WO1991007408A1 - Imidazoquinoxaline compounds, their preparation and use - Google Patents
Imidazoquinoxaline compounds, their preparation and use Download PDFInfo
- Publication number
- WO1991007408A1 WO1991007408A1 PCT/DK1990/000303 DK9000303W WO9107408A1 WO 1991007408 A1 WO1991007408 A1 WO 1991007408A1 DK 9000303 W DK9000303 W DK 9000303W WO 9107408 A1 WO9107408 A1 WO 9107408A1
- Authority
- WO
- WIPO (PCT)
- Prior art keywords
- compound
- general formula
- alkyl
- set forth
- forth above
- Prior art date
- Legal status (The legal status is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the status listed.)
- Ceased
Links
Classifications
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D487/00—Heterocyclic compounds containing nitrogen atoms as the only ring hetero atoms in the condensed system, not provided for by groups C07D451/00 - C07D477/00
- C07D487/02—Heterocyclic compounds containing nitrogen atoms as the only ring hetero atoms in the condensed system, not provided for by groups C07D451/00 - C07D477/00 in which the condensed system contains two hetero rings
- C07D487/04—Ortho-condensed systems
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P25/00—Drugs for disorders of the nervous system
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P25/00—Drugs for disorders of the nervous system
- A61P25/08—Antiepileptics; Anticonvulsants
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P25/00—Drugs for disorders of the nervous system
- A61P25/20—Hypnotics; Sedatives
Definitions
- the present invention relates to therapeutically active imidazoquinoxaline compounds, a method of preparing the same, pharmaceutical compositions comprising the compounds, and to methods of treating therewith.
- the novel compounds are useful in psychopharmaceutical applications, e.g., in the treatment of central nervous system ailments, for example, as anticonvulsants or anxiolytics.
- R 1 and R2 independently are hydrogen, straight
- R is hy ⁇ drogen, straight or branched C 1 _ ⁇ -alkyl, straight or branched C 2 _ ⁇ - alkenyl, or aralkyl or aroylalkyl which may optionally be substituted with halogen or C 1 _,- alkoxy;
- R 4 and R5 independently are hydrogen, halogen,
- the invention also relates to a method of preparing the above mentioned compounds. This method comprises:
- R 3, R4 and R5 have the meanings set forth above, with a compound having the general formula III
- R 1, R3, R4 and R5 have the meanings set forth above, or
- R , R R 3, R4 and R5 have the meanings set forth above.
- the leaving group, Y may be any suitable leaving group and, for example, those disclosed in U.S. Patents 4,031,079 or 4,359,420, for example, halogen, alkyl- thio, e.g., methylthio, aralkylthio, N-nitrosoalkyl- amino, alkoxy, mercapto, -OP(0)(OR) 2 wherein R is lower-alkyl or -OP(0)(NR'R * * ) 2 wherein R * and R * * each represents lower- alkyl or phenyl, or together with the nitrogen atom to which they are attached represent a heterocyclic radical such as morpholino, pyrrolidino, piperidino, or methylpiperazino.
- halogen alkyl- thio, e.g., methylthio, aralkylthio, N-nitrosoalkyl- amino, alkoxy, mercapto, -OP(0)(OR) 2 wherein R is
- the reaction is pre ⁇ ferably carried out under alkaline conditions, i.e., in the presence of a base, and among bases alkali metal, e.g., potassium or sodium, alkoxides or hydrides are preferred.
- the reaction is preferably conducted in the presence of an organic solvent which is nonreactive with the reactants and products of reaction under the conditions of reaction, especially an anhydrous sol ⁇ vent and preferably an anhydrous aprotic solvent such as dimethylformamide (DMF) or the like.
- the temperature range employed may be any range suitable for the reac ⁇ tion to proceed at a reasonable rate and without undue delay or decomposition and a range from a minus forty (-40) degrees Celsius to about room temperature is accordingly usually particularly suitable.
- the starting materials may be prepared from commer ⁇ cially available organic compounds and by using well known synthetic methods.
- the pharmaceutical properties of the compounds of the invention can be illustrated by determining their capability for displacing radioactive labelled fluni- trazepam from benzodiazepine receptors.
- the displacement activity of the compounds of the invention may be found by determining the EDj- n value.
- the ED J - value represents the dose (mg/kg) of a test substance which causes the specific binding of fluni- trazepam to benzodiazepine receptors in a living brain to be reduced to 50% of the control value.
- H-flunitraze- 3 pam H-FNM
- H-FNM H-flunitraze- 3 pam
- H-FNM can be partly or completely prevented by simul ⁇ taneous or prior administration of pharmacologically active benzodiazepines and by some benzodiazepine-like agents (Chang and Snyder, Eur.J. Pharmacol. 48, 212-218 (1978)).
- Test procedure Suspensions of test substances (2 mg/- ml) are prepared in 5% Duphasol-X (TM Duphar, castor oil- ethylene oxide derivative for emulsifying and solubilizing oil and other water-insoluble substances ) by sonification for 10 min using a Branson B15 microtip ultrasonifier (setting 7). Groups of three mice (female, NMR, 18-22 grams) are injected with the test substance at 100 mg/kg intraperitoneally. Fifteen minutes after test substance administration the mice are challenged with 4 ⁇ Ci intravenously of H-FNM (70-90 Ci/mole) in
- H-FNM administration mice are sacrificed by decapita ⁇ tion, the forebrains rapidly excised (within 30 sec) and homogenized in 12 ml of icecold 25 mM KH 2 P0,, pH 7.1, using an Ultra-Turrax homogenizer fitted with an N 10 shaft. Two aliquots of 1 ml are immediately fil ⁇ tered through Whatman GF/C glassfibre filters and wash ⁇ ed with 2 x 5 ml of the above mentioned buffer. The amounts of radioactivity on the filters are determined by conventional scintillation counting. One group of untreated mice serves as control. One to three mice are injected with 25 ⁇ g/kg clonazepam i.p. 30 minutes
- test substances are administered in doses, which are factors of 3.16 times lower than 100 mg/kg.
- the ED,- n for a test sub- stance is defined as that dose which inhibits 50% of 3 specific H-FNM binding.
- Specific binding is the amount of binding in controls minus the amount binding in clonazepam- treated mice.
- the ED 5 _ value is determined from dose re ⁇ sponse curves. If only one dose of test substance is administered the ED_ 0 value is calculated as follows, provided that the inhibition of specific binding is within the range of 25-75%:
- the compound of the invention together with a con- ventional adjuvant, carrier, or diluent, and if desired in the form of a pharmaceutically-acceptable acid addi ⁇ tion salt thereof, may be placed into the form of phar ⁇ maceutical compositions and unit dosages thereof, and in such form may be employed as solids, such as tablets or filled capsules, or liquids, such as solutions, suspensions, emulsions, elixirs, or capsules filled with the same, all for oral use, in the form of suppo ⁇ sitories for rectal administration; or in the form of sterile injectable solutions for parenteral (including subcutaneous) use.
- Such pharmaceutical compositions and unit dosage forms thereof may comprise conventional ingredients in conventional proportions, with or with ⁇ out additional active compounds or principles, and such unit dosage forms may contain any suitable effective central nervous system ailment alleviating amount of the active ingredient commensurate with the intended daily dosage range to be employed.
- Tablets containing one (1) milligram of active ingredient or, more broad ⁇ ly, one (1) to thirty (30) milligrams, per tablet, are accordingly suitable representative unit dosage forms.
- the compounds of this invention can thus be used for the formulation of pharmaceutical preparations, e.g., for oral and parenteral administration to mammals in ⁇ cluding humans, in accordance with conventional methods of galenic pharmacy.
- excipients are such pharmaceutically accept ⁇ able organic or inorganic carrier substances suitable for parenteral or oral application which do not delet ⁇ eriously react with the active compound.
- Such carriers are water, salt solutions, alcohols, polyethylene glycols, polyhydroxyethoxylated castor oil, gelatin, lactose, amylose, magnesium ste- arate, talc, silicic acid, fatty acid monoglycerides and diglycerides, pentaerythritol fatty acid esters, hydroxymethylcellulose and polyvinylpyrrolidone.
- the pharmaceutical preparations can be sterilized and mixed, if desired, with auxilliary agents, such as lubricants, preservatives, stabilizers, wetting agents, emulsifiers, salt for influencing osmotic pressure, buffers and/or coloring substances and the like, which do not deleteriously react with the active compound.
- auxilliary agents such as lubricants, preservatives, stabilizers, wetting agents, emulsifiers, salt for influencing osmotic pressure, buffers and/or coloring substances and the like, which do not deleteriously react with the active compound.
- injectable solutions or suspensions preferably aqueous solutions with the active compound dissolved in polyhydroxylated castor oil.
- Ampoules are convenient unit dosage forms.
- tablets, dragees, or capsules having talc and/or a carbohydrate carrier or binder or the like, the carrier preferably being lactose and/or corn starch and/or po ⁇ tato starch.
- a syrup, elixir or like can be used when a sweetened vehicle can be employed.
- the compounds of the invention are dispensed in unit dosage form comprising 0.05-100 mg in a pharmaceutically-acceptable carrier per unit dosage.
- a typical tablet which may be prepared by conventional tabletting techniques contains:
- the compounds of the invention are extremely useful in the treatment of central nervous system ailments or disorders, when administered in an amount effective for the alleviation, amelioration, or elimination thereof.
- the important CNS activity of the compounds of the invention includes both anticonvulsant and anxiolytic activities along with a low toxicity, together presenting a most favorable therapeutic index.
- the compounds of the invention may accordingly be ad- ministered to a subject, e.g., a living animal or a human body, in need of the same for the treatment, alleviation, amelioration, or elimination of an indi ⁇ cation, associated with the central nervous system and the socalled benzodiazepin receptors, which requires such psychopharmaceutical treatment, e.g., especially convulsion and/or anxiety states, if desired in the form of a pharmaceuticallyacceptable acid addition salt thereof (such as the hydrobromide, hydrochloride, or sulfate, in any event prepared in the usual or conven ⁇ tional manner, e.g., evaporation to dryness of the free base in solution together with the acid), ordina- rily concurrently, simultaneously, or together with a pharmaceutically-acceptable carrier or diluent, espe ⁇ cially and preferably in the form of a pharmaceutical composition thereof, whether by oral, rectal, or paren ⁇ teral (including subcutaneous) route, in
- Suitable dosage ranges are 1-200 milligrams daily, 1-100 milligrams daily, and especially 1-30 milligrams daily, depending as usual upon the exact mode of administration, form in which administered, the indication toward which the administration is directed, the subject involved and the body weight of the subject involved, and the preference and experience of the physician or veterinarian in charge.
Landscapes
- Organic Chemistry (AREA)
- Chemical & Material Sciences (AREA)
- Health & Medical Sciences (AREA)
- Neurosurgery (AREA)
- Life Sciences & Earth Sciences (AREA)
- Neurology (AREA)
- Bioinformatics & Cheminformatics (AREA)
- Engineering & Computer Science (AREA)
- Chemical Kinetics & Catalysis (AREA)
- General Chemical & Material Sciences (AREA)
- Medicinal Chemistry (AREA)
- Nuclear Medicine, Radiotherapy & Molecular Imaging (AREA)
- Veterinary Medicine (AREA)
- Pharmacology & Pharmacy (AREA)
- Biomedical Technology (AREA)
- Animal Behavior & Ethology (AREA)
- General Health & Medical Sciences (AREA)
- Public Health (AREA)
- Anesthesiology (AREA)
- Pain & Pain Management (AREA)
- Pharmaceuticals Containing Other Organic And Inorganic Compounds (AREA)
- Nitrogen Condensed Heterocyclic Rings (AREA)
- Nitrogen And Oxygen Or Sulfur-Condensed Heterocyclic Ring Systems (AREA)
- Agricultural Chemicals And Associated Chemicals (AREA)
Abstract
Description

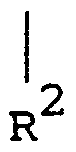


Claims


Priority Applications (5)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| DE69009493T DE69009493T2 (en) | 1989-11-22 | 1990-11-22 | IMIDAZOQUINOXALINE COMPOUNDS, THEIR PRODUCTION AND USE. |
| KR1019920701199A KR920703593A (en) | 1989-11-22 | 1990-11-22 | Imidazoquinoxaline compound, preparation method and use thereof |
| EP90917143A EP0502885B1 (en) | 1989-11-22 | 1990-11-22 | Imidazoquinoxaline compounds, their preparation and use |
| NO921886A NO300064B1 (en) | 1989-11-22 | 1992-05-13 | Analogous Process for the Preparation of Therapeutically Active Imidazoquinoxaline Compounds |
| FI922269A FI97228C (en) | 1989-11-22 | 1992-05-19 | Process for the preparation of novel therapeutically useful 4-oxo-imidazo / 1,5-a / quinoxaline derivatives |
Applications Claiming Priority (2)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| DK588389A DK588389D0 (en) | 1989-11-22 | 1989-11-22 | HETEROCYCLIC COMPOUNDS, THEIR PREPARATION AND USE |
| DK5883/89 | 1989-11-22 |
Publications (1)
| Publication Number | Publication Date |
|---|---|
| WO1991007408A1 true WO1991007408A1 (en) | 1991-05-30 |
Family
ID=8145913
Family Applications (1)
| Application Number | Title | Priority Date | Filing Date |
|---|---|---|---|
| PCT/DK1990/000303 Ceased WO1991007408A1 (en) | 1989-11-22 | 1990-11-22 | Imidazoquinoxaline compounds, their preparation and use |
Country Status (19)
| Country | Link |
|---|---|
| US (1) | US5102885A (en) |
| EP (1) | EP0502885B1 (en) |
| JP (1) | JPH05503084A (en) |
| KR (1) | KR920703593A (en) |
| AT (1) | ATE106405T1 (en) |
| AU (1) | AU645508B2 (en) |
| CA (1) | CA2073046A1 (en) |
| DE (1) | DE69009493T2 (en) |
| DK (2) | DK588389D0 (en) |
| ES (1) | ES2055454T3 (en) |
| FI (1) | FI97228C (en) |
| HU (1) | HUT63420A (en) |
| IE (1) | IE69960B1 (en) |
| IL (1) | IL96372A (en) |
| NO (1) | NO300064B1 (en) |
| NZ (1) | NZ236156A (en) |
| PT (1) | PT95972B (en) |
| WO (1) | WO1991007408A1 (en) |
| ZA (1) | ZA909304B (en) |
Cited By (1)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| FR2722789A1 (en) * | 1994-07-20 | 1996-01-26 | Rhone Poulenc Rorer Sa | IMIDAZO (1,2-A) INDENO (1,2-E) PYRAZINE-2-CARBOXYLIC ACID DERIVATIVES, THEIR PREPARATION AND THE MEDICINAL PRODUCTS CONTAINING THE SAME |
Families Citing this family (1)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| DK588489D0 (en) * | 1989-11-22 | 1989-11-22 | Ferrosan As | HETEROCYCLIC COMPOUNDS, THEIR PREPARATION AND USE |
Citations (3)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| EP0225013A1 (en) * | 1985-10-17 | 1987-06-10 | A/S Ferrosan | Heterocyclic compounds and their preparation and use |
| EP0283162A2 (en) * | 1987-03-18 | 1988-09-21 | Novo Nordisk A/S | Heterocyclic compounds and their preparation and use |
| EP0320136A2 (en) * | 1987-12-08 | 1989-06-14 | Novo Nordisk A/S | Imidazoquinoxaline compounds and their preparation and use |
Family Cites Families (1)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| FR2659329B1 (en) * | 1990-03-09 | 1994-06-03 | Adir | NOVEL IMIDAZO [1,2-C] QUINAZOLINE DERIVATIVES, THEIR PREPARATION PROCESS AND THE PHARMACEUTICAL COMPOSITIONS CONTAINING THEM. |
-
1989
- 1989-11-22 DK DK588389A patent/DK588389D0/en not_active Application Discontinuation
-
1990
- 1990-11-16 IL IL9637290A patent/IL96372A/en not_active IP Right Cessation
- 1990-11-19 IE IE415790A patent/IE69960B1/en not_active IP Right Cessation
- 1990-11-20 US US07/616,136 patent/US5102885A/en not_active Expired - Fee Related
- 1990-11-20 ZA ZA909304A patent/ZA909304B/en unknown
- 1990-11-20 NZ NZ236156A patent/NZ236156A/en unknown
- 1990-11-22 DK DK90917143.1T patent/DK0502885T3/en active
- 1990-11-22 CA CA002073046A patent/CA2073046A1/en not_active Abandoned
- 1990-11-22 KR KR1019920701199A patent/KR920703593A/en not_active Ceased
- 1990-11-22 HU HU9201709A patent/HUT63420A/en unknown
- 1990-11-22 ES ES90917143T patent/ES2055454T3/en not_active Expired - Lifetime
- 1990-11-22 PT PT95972A patent/PT95972B/en not_active IP Right Cessation
- 1990-11-22 WO PCT/DK1990/000303 patent/WO1991007408A1/en not_active Ceased
- 1990-11-22 AU AU67429/90A patent/AU645508B2/en not_active Ceased
- 1990-11-22 DE DE69009493T patent/DE69009493T2/en not_active Expired - Fee Related
- 1990-11-22 AT AT90917143T patent/ATE106405T1/en not_active IP Right Cessation
- 1990-11-22 EP EP90917143A patent/EP0502885B1/en not_active Expired - Lifetime
- 1990-11-22 JP JP3500074A patent/JPH05503084A/en active Pending
-
1992
- 1992-05-13 NO NO921886A patent/NO300064B1/en unknown
- 1992-05-19 FI FI922269A patent/FI97228C/en not_active IP Right Cessation
Patent Citations (3)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| EP0225013A1 (en) * | 1985-10-17 | 1987-06-10 | A/S Ferrosan | Heterocyclic compounds and their preparation and use |
| EP0283162A2 (en) * | 1987-03-18 | 1988-09-21 | Novo Nordisk A/S | Heterocyclic compounds and their preparation and use |
| EP0320136A2 (en) * | 1987-12-08 | 1989-06-14 | Novo Nordisk A/S | Imidazoquinoxaline compounds and their preparation and use |
Cited By (2)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| FR2722789A1 (en) * | 1994-07-20 | 1996-01-26 | Rhone Poulenc Rorer Sa | IMIDAZO (1,2-A) INDENO (1,2-E) PYRAZINE-2-CARBOXYLIC ACID DERIVATIVES, THEIR PREPARATION AND THE MEDICINAL PRODUCTS CONTAINING THE SAME |
| WO1996002544A1 (en) * | 1994-07-20 | 1996-02-01 | Rhone-Poulenc Rorer S.A. | IMIDAZO[1,2-a]INDENO[1,2-e]PYRAZINE-2-CARBOXYLIC ACID DERIVATIVES, PREPARATION THEREOF AND DRUGS CONTAINING SAME |
Also Published As
| Publication number | Publication date |
|---|---|
| KR920703593A (en) | 1992-12-18 |
| PT95972B (en) | 1998-01-30 |
| JPH05503084A (en) | 1993-05-27 |
| IL96372A0 (en) | 1991-08-16 |
| FI922269L (en) | 1992-05-19 |
| FI97228C (en) | 1996-11-11 |
| NZ236156A (en) | 1993-01-27 |
| NO300064B1 (en) | 1997-04-01 |
| DE69009493T2 (en) | 1994-12-08 |
| IE904157A1 (en) | 1991-05-22 |
| NO921886L (en) | 1992-05-13 |
| AU645508B2 (en) | 1994-01-20 |
| CA2073046A1 (en) | 1991-05-23 |
| ATE106405T1 (en) | 1994-06-15 |
| DE69009493D1 (en) | 1994-07-07 |
| PT95972A (en) | 1991-09-13 |
| IL96372A (en) | 1994-10-21 |
| DK588389D0 (en) | 1989-11-22 |
| US5102885A (en) | 1992-04-07 |
| EP0502885A1 (en) | 1992-09-16 |
| AU6742990A (en) | 1991-06-13 |
| DK0502885T3 (en) | 1994-09-26 |
| IE69960B1 (en) | 1996-10-16 |
| HU9201709D0 (en) | 1992-09-28 |
| ES2055454T3 (en) | 1994-08-16 |
| FI922269A0 (en) | 1992-05-19 |
| EP0502885B1 (en) | 1994-06-01 |
| ZA909304B (en) | 1991-09-25 |
| HUT63420A (en) | 1993-08-30 |
| FI97228B (en) | 1996-07-31 |
| NO921886D0 (en) | 1992-05-13 |
Similar Documents
| Publication | Publication Date | Title |
|---|---|---|
| US4774245A (en) | Imidazoquinoxaline compounds | |
| US5034530A (en) | Imidazoquinoxaline compounds and their preparation and use | |
| EP0241682B1 (en) | Novel benzodiazepine derivatives and their preparation and use | |
| US4999354A (en) | Imidazoquinoxaline compounds and their preparation and use | |
| AU673814B2 (en) | Imidazoquinazoline compounds and their preparation and use | |
| EP0226282B1 (en) | Heterocyclic compounds and their preparation and use | |
| EP0201678B1 (en) | 3-(5-cyclopropyl-1,2,4-oxadiazol-3-yl)-5,6-dihydro-5-methyl-6-oxo-4h-imidazo(1,5-a)(1,4)benzodiazepine, a process for its preparation and pharmaceutical compositions | |
| US5075304A (en) | Imidazoquinoxaline compounds and their preparation and use | |
| AU645508B2 (en) | Imidazoquinoxaline compounds, their preparation and use | |
| EP0502038B1 (en) | Imidazoquinoxaline compounds, their preparation and use | |
| AU8063391A (en) | Heterocyclic compounds and their preparation and use | |
| EP0202441B1 (en) | Oxadiazolylimidazobenzodiazepines, process for their preparation and pharmaceutical compositions | |
| US4745112A (en) | Oxadiazolylimidazobenzodiazepine, compositions, and method III | |
| AU650733B2 (en) | Tetracyclic imidazoquinazoline derivatives, process for their preparation and pharmaceutical compositions containing them |
Legal Events
| Date | Code | Title | Description |
|---|---|---|---|
| AK | Designated states |
Kind code of ref document: A1 Designated state(s): AU CA FI HU JP KR NO |
|
| AL | Designated countries for regional patents |
Kind code of ref document: A1 Designated state(s): AT BE CH DE DK ES FR GB GR IT LU NL SE |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 2073046 Country of ref document: CA |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 922269 Country of ref document: FI |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 1990917143 Country of ref document: EP |
|
| WWP | Wipo information: published in national office |
Ref document number: 1990917143 Country of ref document: EP |
|
| WWG | Wipo information: grant in national office |
Ref document number: 1990917143 Country of ref document: EP |
|
| WWG | Wipo information: grant in national office |
Ref document number: 922269 Country of ref document: FI |