WO1994002493A1 - Phosphonated arylethanolamine compounds with anti-hyperglycemic and/or anti-obesity activity - Google Patents

Phosphonated arylethanolamine compounds with anti-hyperglycemic and/or anti-obesity activity Download PDF

Info

Publication number
WO1994002493A1
WO1994002493A1 PCT/GB1993/001554 GB9301554W WO9402493A1 WO 1994002493 A1 WO1994002493 A1 WO 1994002493A1 GB 9301554 W GB9301554 W GB 9301554W WO 9402493 A1 WO9402493 A1 WO 9402493A1
Authority
WO
WIPO (PCT)
Prior art keywords
formula
compound
pharmaceutically acceptable
ester
amide
Prior art date
Legal status (The legal status is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the status listed.)
Ceased
Application number
PCT/GB1993/001554
Other languages
French (fr)
Inventor
Lee James Beeley
John Michael Berge
Richard Lewis Jarvest
Current Assignee (The listed assignees may be inaccurate. Google has not performed a legal analysis and makes no representation or warranty as to the accuracy of the list.)
SmithKline Beecham Ltd
Original Assignee
SmithKline Beecham Ltd
Priority date (The priority date is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the date listed.)
Filing date
Publication date
Application filed by SmithKline Beecham Ltd filed Critical SmithKline Beecham Ltd
Priority to DE69311118T priority Critical patent/DE69311118T2/en
Priority to EP93917898A priority patent/EP0652884B1/en
Priority to JP6504304A priority patent/JPH07509458A/en
Priority to AU47151/93A priority patent/AU4715193A/en
Publication of WO1994002493A1 publication Critical patent/WO1994002493A1/en
Anticipated expiration legal-status Critical
Ceased legal-status Critical Current

Links

Classifications

    • C—CHEMISTRY; METALLURGY
    • C07—ORGANIC CHEMISTRY
    • C07F—ACYCLIC, CARBOCYCLIC OR HETEROCYCLIC COMPOUNDS CONTAINING ELEMENTS OTHER THAN CARBON, HYDROGEN, HALOGEN, OXYGEN, NITROGEN, SULFUR, SELENIUM OR TELLURIUM
    • C07F9/00—Compounds containing elements of Groups 5 or 15 of the Periodic Table
    • C07F9/02—Phosphorus compounds
    • C07F9/547—Heterocyclic compounds, e.g. containing phosphorus as a ring hetero atom
    • C07F9/655—Heterocyclic compounds, e.g. containing phosphorus as a ring hetero atom having oxygen atoms, with or without sulfur, selenium, or tellurium atoms, as the only ring hetero atoms
    • C07F9/65515—Heterocyclic compounds, e.g. containing phosphorus as a ring hetero atom having oxygen atoms, with or without sulfur, selenium, or tellurium atoms, as the only ring hetero atoms the oxygen atom being part of a five-membered ring
    • A—HUMAN NECESSITIES
    • A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
    • A61P1/00—Drugs for disorders of the alimentary tract or the digestive system
    • A—HUMAN NECESSITIES
    • A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
    • A61P15/00—Drugs for genital or sexual disorders; Contraceptives
    • A—HUMAN NECESSITIES
    • A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
    • A61P3/00—Drugs for disorders of the metabolism
    • A61P3/04—Anorexiants; Antiobesity agents
    • A—HUMAN NECESSITIES
    • A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
    • A61P3/00—Drugs for disorders of the metabolism
    • A61P3/08—Drugs for disorders of the metabolism for glucose homeostasis
    • A—HUMAN NECESSITIES
    • A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
    • A61P3/00—Drugs for disorders of the metabolism
    • A61P3/08—Drugs for disorders of the metabolism for glucose homeostasis
    • A61P3/10—Drugs for disorders of the metabolism for glucose homeostasis for hyperglycaemia, e.g. antidiabetics
    • A—HUMAN NECESSITIES
    • A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
    • A61P9/00—Drugs for disorders of the cardiovascular system
    • A61P9/10—Drugs for disorders of the cardiovascular system for treating ischaemic or atherosclerotic diseases, e.g. antianginal drugs, coronary vasodilators, drugs for myocardial infarction, retinopathy, cerebrovascula insufficiency, renal arteriosclerosis
    • C—CHEMISTRY; METALLURGY
    • C07—ORGANIC CHEMISTRY
    • C07F—ACYCLIC, CARBOCYCLIC OR HETEROCYCLIC COMPOUNDS CONTAINING ELEMENTS OTHER THAN CARBON, HYDROGEN, HALOGEN, OXYGEN, NITROGEN, SULFUR, SELENIUM OR TELLURIUM
    • C07F9/00—Compounds containing elements of Groups 5 or 15 of the Periodic Table
    • C07F9/02—Phosphorus compounds
    • C07F9/28—Phosphorus compounds with one or more P—C bonds
    • C07F9/38—Phosphonic acids [RP(=O)(OH)2]; Thiophosphonic acids ; [RP(=X1)(X2H)2(X1, X2 are each independently O, S or Se)]
    • C07F9/3804—Phosphonic acids [RP(=O)(OH)2]; Thiophosphonic acids ; [RP(=X1)(X2H)2(X1, X2 are each independently O, S or Se)] not used, see subgroups
    • C07F9/3808—Acyclic saturated acids which can have further substituents on alkyl
    • C—CHEMISTRY; METALLURGY
    • C07—ORGANIC CHEMISTRY
    • C07F—ACYCLIC, CARBOCYCLIC OR HETEROCYCLIC COMPOUNDS CONTAINING ELEMENTS OTHER THAN CARBON, HYDROGEN, HALOGEN, OXYGEN, NITROGEN, SULFUR, SELENIUM OR TELLURIUM
    • C07F9/00—Compounds containing elements of Groups 5 or 15 of the Periodic Table
    • C07F9/02—Phosphorus compounds
    • C07F9/28—Phosphorus compounds with one or more P—C bonds
    • C07F9/38—Phosphonic acids [RP(=O)(OH)2]; Thiophosphonic acids ; [RP(=X1)(X2H)2(X1, X2 are each independently O, S or Se)]
    • C07F9/40—Esters thereof
    • C07F9/4003—Esters thereof the acid moiety containing a substituent or a structure which is considered as characteristic
    • C07F9/4006—Esters of acyclic acids which can have further substituents on alkyl

Definitions

  • This invention relates to novel compounds, to a process for preparing such compounds, to pharmaceutical compositions containing such compounds and to the use of such compounds and compositions in medicine and agriculture.
  • European patent application, publication number 0455006 discloses certain arylethanolamine derivatives which are stated to possess anti-diabetic and/or anti- hyperglycaemic and/or anti-obesity activity. It has been discovered that a series of novel phosphonated arylethanolamine derivatives have good ⁇ -adrenoreceptor agonist activity and in particular show good selectivity for ⁇ 3-adrenoreceptors versus ⁇ i- or ⁇ 2-adrenoreceptors. These compounds arc therefore indicated to have good anti-hyperglycaemic and/or anti- obesity activity coupled with especially good selectivity from cardiac and tremorigenic side effects.
  • the present invention provides a compound of formula (I):
  • R° represents a substituted or unsubstituted aryl group
  • R.1 represents hydrogen or an alkyl group
  • R- represents a moiety of formula (a):
  • R ⁇ and R ⁇ each independently represent hydrogen, alkyl, hydroxyalkyl, cycloalkyl or R ⁇ together with R ⁇ represents (CH2)n wherein n is 2, 3 or 4; T represents hydrogen, nitrile or a group -CO.R s wherein R s represents hydroxy, alkoxy or a group -NR l R u wherein R and R u independently represent hydrogen, alkyl or R l and R u together with the nitrogen to which they are attached represent a saturated heterocyclic group; and
  • R- represents hydrogen, halogen, alkyl or alkoxy.
  • R° represents a phenyl group optionally substituted with halogen or a hydroxy
  • examples include chlorophenyl, especially 3-chlorophenyl, and hydroxyphenyl, especially 3,4-bis-hydroxyphenyl.
  • R* is an alkyl group.
  • R! is alkyl
  • it is favourably a C ⁇ .g alkyl group, especially a methyl group.
  • R ⁇ and R-5 each independently represent alkyl, for example ethyl propyl, or butyl.
  • R ⁇ is alkyl, for example ethyl, propyl or butyl, especially ethyl, and R- is hydrogen.
  • R- represents hydrogen
  • T represents hydrogen or a group -CO.R s wherein R s is hydroxy or alkoxy.
  • T represents hydrogen.
  • the compounds of formula (I) have one or two asymmetric carbon atoms, marked with an asterisk (*) or two asterisks (**) in the formula. These compounds may therefore exist in up to four stereoisomeric forms.
  • the present invention encompasses all stereoisomers of the compounds of the general formula (I) whether free from other isomers, or admixed with other isomers in any proportion, such as mixtures of diastereoisomers and racemic mixtures of enantiomers.
  • the asymmetric carbon atom indicated by a single asterisk (*) is in the R-configuration.
  • the asymmetric carbon atom indicated by two asterisks (**) is in the R-configuration.
  • One suitable form of a compound of formula (I) is a diastereomeric mixture of the RR and SS enantiomers.
  • a compound of formula (I) is the RR enantiomer.
  • the term 'alkyl' when used alone or when forming part of other groups (such as the 'alkoxy' group) includes straight- or branched-chain alkyl groups containing 1 to 12 carbon atoms, suitably 1 to 6 carbon atoms, examples include methyl, ethyl, n-propyl, isopropyl, n-butyl, isobutyl or tert-butyl group.
  • 'cycloalkyl' includes C3_ cycloalkyl groups, especially C5 or Cg cycloalkyl groups.
  • halogen refers to fluorine, chlorine, bromine and iodine, preferably chlorine.
  • 'aryl' includes phenyl and naphthyl optionally substituted with up to five, preferably up to three, groups selected from halogen, alkyl, phenyl, alkoxy, haloalkyl, hydroxyalkyl, hydroxy, amino, nitro, carboxy and pharmaceutically acceptable salts, esters and amides thereof, alkoxycarbonylalkyl, alkylcarbonyloxy and alkylcarbonyl groups.
  • a preferred aryl group is a substituted or unsubstituted phenyl group.
  • Preferred optional substituents for the aryl group include up to three substituents selected from halogen, hydroxy, alkoxy, hydroxyalkyl and amino.
  • Suitable pharmaceutically acceptable esters of carboxyl groups include alkyl esters, especially C ⁇ _6 alkyl esters such as methyl.
  • Suitable pharmaceutically acceptable amides are those of formula -CONR x Ry wherein R " and RY each independently represent hydrogen, alkyl or alkoxyalkyl.
  • Suitable pharmaceutically acceptable salts include acid addition salts, salts of carboxy groups and salts of phosphonic acid groups.
  • Suitable pharmaceutically acceptable acid addition salts include salts with inorganic acids such, for example, as hydrochloric acid, hydrobromic acid, orthophosphoric acid or sulphuric acid, or with organic acids such, for example as methanesulphonic acid, toluenesulophonic acid, acetic acid, propionic acid, lactic acid, citric acid, fumaric acid, malic acid, succinic acid, salicylic acid, maleic acid or acetylsalicylic acid.
  • inorganic acids such, for example, as hydrochloric acid, hydrobromic acid, orthophosphoric acid or sulphuric acid
  • organic acids such, for example as methanesulphonic acid, toluenesulophonic acid, acetic acid, propionic acid, lactic acid, citric acid, fumaric acid, malic acid, succinic acid, salicylic acid, maleic acid or acetylsalicylic acid.
  • Suitable pharmaceutically acceptable salts of carboxy groups or phosphonic acid groups include metal salts, such as for example aluminium, alkali metal salts such as sodium or potassium, alkaline earth metal salts such as calcium or magnesium and ammonium or substituted ammonium salts, for example those with C g alkylamines such as triethylamine, hydroxy-C ⁇ _6 alkylamines such as 2-hydroxyethylamine, bis-(2- hydroxyethyl)-amine or tri-(2-hydroxyethyl)-amine, cycloalkylamines such as bicyclohexylamine, or with procaine, dibenzylpiperidine, N-benzyl- ⁇ -phenethylamine, dehydroabietylamine, N,N'-bisdehydroabietylamine, glucamine, N-methylglucamine or bases of the pyridine type such as pyridine, collidine or quinoline.
  • metal salts such as for example aluminium, alkal
  • Suitable pharmaceutically acceptable solvates are conventional solvates, preferably hydrates.
  • the invention also provides a process for the preparation of a compound of formula (I), or a pharmaceutically acceptable salt, ester or amide thereof or a pharmaceutically acceptable solvate thereof, which process comprises reducing a compound of formula (II):
  • R°, Rl, R2 and R ⁇ are as defined in relation to formula (I), and thereafter, if necessary, carrying out one or more of the following optional steps:
  • the reduction of the compound of formula (II) may be carried out using any suitable reduction procedure, for example by using catalytic reduction in the presence of hydrogen.
  • Suitable catalysts include platinum oxide or 10% palladium on charcaol.
  • Suitable reduction conditions include using an alkanolic solvent such as methanol, at any temperature providing a convenient rate of formation of the required product, for example when using the platinum catalyst the reaction may conveniently be carried out at ambient temperature or when using the palladium catalyst the reaction may be carried out at a medium temperature such as 50°C, under a pressure of 1-5 atmospheres of hydrogen.
  • an alkanolic solvent such as methanol
  • the compound of formula (II) may be prepared by reacting a compound of formula (III): OH o l
  • R2 and R ⁇ are as defined in relation to formula (I).
  • the reaction between compounds of formulae (III) and (IN) may be carried out under conventional animation conditions, for example in a solvent such as methanol or toluene.
  • the compound of formula (II) is prepared in-situ by reacting a compound of the above defined formula (III) with a compound of the above defined formula (IV) under reductive amination conditions.
  • Suitable reductive amination conditions include reacting the compounds of formula (III) and (IV) in an alkanolic solvent, such as methanol, in the presence of a suitable reduction catalyst.
  • a compound of formula (IV) may be prepared by reducing a compound of formula (V):
  • R , R-- and R ⁇ are as defined in relation to formula (I).
  • the reduction of the compound of formula (V) may conveniendy be carried out using iron powder in the presence of acetic acid in an aqueous solvent such as aqueous methanol, at any temperature providing a suitable rate of formation of the required product, generally at an elevated temperature and conveniently at the reflux temperature of the solvent.
  • an aqueous solvent such as aqueous methanol
  • a compound of formula (V) may be prepared by reacting a compound of formula (VI):
  • R-- and R3 are as defined in relation to formula (I), with nitroalkane.
  • the carbon atom of the -CHO group of compound of formula (VI) is in an activated form, a suitable activated form being provided by forming an imine of the said carbonyl group:
  • the imine may be prepared by reacting the compound of formula (VI) with an amine, suitably a primary alkyl amine such as n-butylamine.
  • reaction of the compound of formula (VI) and the amine may be carried out in any suitable solvent, such as toluene, at any temperature providing a suitable rate of formation of the required product, generally at an elevated temperature such as the reflux temperature of the solvent; and preferably in the presence of a catalytic amount of toluenesulphonic acid.
  • suitable solvent such as toluene
  • the reaction between the compound of formula (VI), and when it is in the form of an imine, and nitroalkane may be carried out in glacial acetic acid, preferably in the presence of an ammonium acetate catalyst, generally at an elevated temperature such as in the range of from 60°C to 120°C, for example 100°C.
  • a compound of formula (VI), wherein T is hydrogen, may be prepared from a compound of formula (Nil):
  • R ⁇ is as defined in relation to formula (I) and L* is a leaving group or atom, generally a fluorine atom, with an activated form of a compound of formula (VIII): HOCH 2 -P-OR 4
  • a suitable activated form of a compound of formula (VIII) is an ionic form, such as a salted form, for example an alkali metal salted form.
  • An activated form of a compound of formula (VIII) may be prepared by use of the appropriate conventional procedure, for example a salted form may be prepared by treating the compound of formula (VHT) with a base such as an alkali metal hydride, for example sodium hydride.
  • a base such as an alkali metal hydride, for example sodium hydride.
  • reaction between the compounds of formulae (VII) and (VIII) may be carried out in any suitable solvent, generally an aprotic solvent such as dimethylformamide or N-methylpyrrolidinone at a low to ambient temperature, for example in the range of from -15°C to 20°C, such as 5°C.
  • aprotic solvent such as dimethylformamide or N-methylpyrrolidinone
  • a compound of formula (IV) may also be prepared by the reaction of a compound of formula (IX):
  • R , R ⁇ and R- are as defined in relation to formula (I) and J represents an oxo group or a protected oxo group, for example an ethan-l,2-dioxy group, with a compound of formula (X):
  • R ⁇ , R ⁇ and T are as defined in relation to formula (I) in the presence of a rhodium salt , and thereafter, as required, removing any protecting group.
  • a suitable rhodium salt is rhodium acetate.
  • the reaction between the compound of formula (IX) and the compound of formula (X) may be carried out under conventional carbene insertion conditions, in an inert solvent, such as benzene at any temperature providing a convenient rate of formation of the required product, generally at an elevated temperature, such as the reflux temperature of the solven; preferably the reaction is carried out under an inert atmosphere such as nitrogen.
  • an inert solvent such as benzene
  • J is a protected oxo group such as an ethan-l,2-dioxy group it may be removed by treatment with an aqueous acid, such as aqueous hydrochloric acid.
  • a compound of formula (I) wherein either the *CH2 or the **CH2 carbon is a chiral carbon may be prepared using conventional stereoselective chemistry from appropriate starting materialsis.
  • One process for the preparation of a compound of formula (I) wherein both the *CH2 and the **CH2 carbon are chiral carbon atoms, or a pharmaceutically acceptable salt, ester or amide thereof or a pharmaceutically acceptable solvate thereof, comprises reacting a compound of formula (XI):
  • Rl, R- ⁇ and R- are as defined in relation to formula (I); and thereafter, if necessary, carrying out one or more of the following optional steps:
  • both the *CH2 and the **CH2 carbon atoms have the R stereochemistry.
  • Suitable reaction conditions for the reaction between the compounds of formulae (XI) and (XII) include using an aprotic solvent such as dimethylsulphoxide, at any temperature providing a convenient rate of formation of the required product, for example at an elevated temperature such as in the range of from 30°C to 100°C, for example 75°C: preferably the reaction is carried out under an inert atmosphere such as nitrogen.
  • a compound of formula (XII) may be prepared by the hydrogenolysis of a compound of formula (XIII):
  • R 1 , R-- and R- are as defined in relation to formula G) and the **CH carbon and ***CH carbon atoms are chiral carbon atoms.
  • catalytic hydrogenolysis is used, using for example 10% palladium on charcaol in the presence of ammonium formate, suitably in an alkanolic solvent such as methanol, at any temperature providing a convenient rate of formation of the required product, for example at ambient temperature; preferably the reaction is carried out in an inert atmosphere, generably under nitrogen .
  • the compound of formula (XIII) may be prepared by stereoselective reduction of a compound of formula (XTV):
  • R 1 , R ⁇ and R- ⁇ are as defined in relation to formula (I) and the ***CH carbon is a chiral carbon.
  • the reduction of the compound of formula (XTV) may be carried out using catalytic reduction in the presence of hydrogen.
  • a preferred catalyst is platinum oxide.
  • Suitable reduction conditions include using an alkanol solvent such as methanol or ethanol, at any temperature providing a convenient rate of formation of the required product, conveniently at ambient temperature using a pressure of 1-5 atmospheres of hydrogen.
  • an alkanol solvent such as methanol or ethanol
  • the compound of formula (XIV) may be prepared by reacting a compound of the above defined formula (IV) with R- ⁇ -methylbenzylamine.
  • the reaction between compounds of formulae (IV) and R- ⁇ - methylbenzylamine may be carried out under conventional amination conditions, for example those described above for the reaction between the compounds of formula (HI) and (TV).
  • the compound of formula (XIV) is prepared inisiflL y reacting a compound of the above defined formula (TV) with R- ⁇ -methylbenzylamine under reductive amination conditions as described in relation to the compounds of formulae (III) and (TV).
  • Suitable conversions of one compound of formula (I) into another compound of formula (I) include converting one group OR 4 into another group OR 4 and/or converting one group OR ⁇ into another group OR*.
  • Suitable conversions of one group OR 4 into another group OR 4 include:
  • the abovementioned conversion (i) may be carried out under conventional phosphonate alkylation methods, using for example the appropriate alcohol (R 4 OH) in the presence of hydrogen chloride.
  • the abovementioned conversion (ii) may be carried out using conventional phosphonate hydrolysis methods, for example by treating the appropriate compound of formula (I) with an alkaline metal hydroxide, such as sodium or lithium hydroxide.
  • an alkaline metal hydroxide such as sodium or lithium hydroxide.
  • This reaction is especially useful when converting OR 4 alkoxy into OR 4 as hydroxy while leaving OR-5 as alkoxy.
  • acidic hydrolysis may be used using for example hydrochloric acid.
  • the abovementioned conversion (iii) may be carried out by first converting OR 4 as alkoxy into OR 4 as hydroxy using the conditions set out in respect of the abovementioned conversion (ii), followed by converting the hydroxy group so formed into another alkoxy group, using the conditions set out in respect of the abovementioned conversion (i).
  • Suitable conversions of one group OR ⁇ into another group OR- ⁇ include analogous conversions to those mentioned above in regard to converting one group OR 4 into another group OR 4 .
  • the protection of any reactive group or atom such as the variables R , R , R2 and R- may be carried out at any appropriate stage in the aforementioned processes.
  • Suitable protecting groups include those used conventionally in the art for the particular group or atom being protected.
  • a keto group may be protected as an ketal.
  • Protecting groups may be prepared and removed using die appropriate conventional procedure.
  • a keto group may be protected as an ketal by reaction with an appropriate alcohol, for example ethan-l,2-diol, in the presence of an acid catalyst, such as 4-toluenesulpho ⁇ ic acid.
  • the said ketal may be removed by use of acidic hydrolysis, for example treatment with hydrochloric acid.
  • a leaving group or atom is any group or atom that will, under the reaction conditions, cleave from the starting material, thus promoting reaction at a specified site. Suitable examples of such groups unless othewise specified are halogen atoms, mesyloxy groups and tosyloxy groups.
  • the compounds of formulae (ID) and (VIII) are known compounds or they may be prepared by processes analogous to those used to prepare known compounds, for example the compounds of formula (LIT) may be prepared according to methods disclosed in J.Org.Chem., 1974, 3_9_, 914; and the compounds of formula (VIII) may be prepared according to methods disclosed in Phosphorus and Sulphur, 1978, ⁇ , 455.
  • the compounds of formula (IX) are known compounds or they may be prepared by methods analogous to those used to prepare known compounds, for example those methods disclosed in European application, publication number 0052963 and United Kingdom patent number 2084577.
  • the compounds of formulae (X) are either known commercially available compounds or they may be prepared from such compounds by conventional procedures for example those disclosed in the Chem. Berichte 1968, 101. 3734 and The Chemistry of Phosphorus, (J. Emsley and D. Hall), Published by Harper and Row, London, 1976.
  • T represents -CO.R l R u
  • T is carboxyl or an ethyl ester thereof, by conventional amination reactions.
  • T is a nitrile group
  • P(OR)3 wherein R is alkyl, suitably ethyl, and thereafter interconverting R into R 4 and or R-5 as required.
  • the compounds of formula (XI) are known compounds or they may be prepared by methods analogous to those used to prepare known compounds, for example those methods disclosed in European application, publication number 0262785.
  • salts, esters, amides and solvates of the compounds mentioned herein may be produced by methods conventional in the art: For example, salts may be prepared by treating a compound of formula (I) with the appropriate acid.
  • Esters of carboxylic acids may be prepared by conventional esterification procedures, for example alkyl esters may be prepared by treating the required carboxylic acid with the appropriate alkanol, generally under acidic conditions.
  • Amides may be prepared using conventional amidation procedures, for example amides of formula CONR s R l may be prepared by treating the relevant carboxylic acid with an amine of formula HNR S R ⁇ wherein R s and R- are as defined above.
  • a C ⁇ . alkyl ester such as a methyl ester of the acid may be treated with an amine of the above defined formula HNR s R l to provide the required amide.
  • mixtures of isomers of the compounds of d e invention may be separated into individual stereoisomers by conventional means, for example by the use of an optically active acid as a resolving agent Suitable optically active acids which maybe used as resolving agents are described in Topics in Stereochemistry', Vol. 6, Wiley Interscience, 1971, Allinger, N.L. and Eliel, W.L. Eds.
  • any enantiomer of a compound of die invention may be obtained by stereospecific synthesis using optically pure starting materials of known configuration.
  • the absolute configuration of compounds may be determined by conventional X-ray crystallographic techniques.
  • the compounds of the present invention have valuable pharmacological properties:
  • the present invention accordingly provides a compound of formula (I) or a pharmaceutically acceptable salt, ester or amide thereof, or a pharmaceutically acceptable solvate thereof, for use as an active therapeutic substance.
  • the present invention provides a compound of formula G)» or a pharmaceutically acceptable salt, ester or amide U ereof thereof, or a pharmaceutically acceptable solvate thereof, for use in the treatment of hyperglycaemia in human or non-human animals.
  • the present invention further provides a compound of formula (I), or pharmaceutically acceptable salt, ester or amide thereof, or a pharmaceutically acceptable solvate thereof, for use in the treatment of obesity in human or non-human animals.
  • a compound of formula (I), or a pharmaceutically acceptable salt, ester or amide thereof, or a pharmaceutically acceptable solvate thereof, may be administered ⁇ £i £g or, preferably, as a pharmaceutical composition also comprising a pharmaceutically acceptable carrier.
  • the present invention also provides a pharmaceutical composition
  • a pharmaceutical composition comprising a compound of formula (I), or a pharmaceutically acceptable salt, ester or amide thereof thereof, or a pharmaceutically acceptable solvate thereof, and a pharmaceutically acceptable carrier therefor.
  • pharmaceutically acceptable embraces compounds, compositions and ingredients for both human and veterinary use: for example the term “pharmaceutically acceptable salt” embraces a veterinarily acceptable salt
  • the composition may, if desired, be in the form of a pack accompanied by written or printed instructions for use.
  • compositions of the present invention will be adapted for oral administration, although compositions for administration by other routes, such as by injection, are also envisaged.
  • Particularly suitable compositions for oral administration are unit dosage forms such as tablets and capsules.
  • Other fixed unit dosage forms, such as powders presented in sachets, may also be used.
  • the carrier may comprise a diluent, filler, disintegrant, wetting agent, lubricant, colourant, flavourant or other conventional adjuvant.
  • Typical carriers include, for example, microcrystalline cellulose, starch, sodium starch glycollate, polyvinylpyrrolidone, polyvinylpolypyrrolidone, magnesium stearate or sodium lauryl sulphate.
  • composition will be formulated in unit dose form.
  • unit dose will normally contain an amount of the active ingredient in d e range of from 0.1 to 1000 mg, more usually 2-100 mg or 0.1 to 500 mg, and more especially 0.1 to 250 mg.
  • the present invention further provides a method for treating hyperglycaemia in a human or non-human mammal, which comprises administering an effective, non-toxic, amount of a compound of formula (I), or a pharmaceutically acceptable salt, ester or amide thereof thereof, or a pharmaceutically acceptable solvate thereof, to a hyperglycaemic human or non-human mammal in need tiiereof.
  • the present invention further provides a method for treating obesity or for the treatment and or prophylaxis of atherosclerosis in a human or non-human mammal, which comprises administering an effective, non-toxic, amount of a compound of formula (I), or a pharmaceutically acceptable salt, ester or amide thereof thereof, or a pharmaceutically acceptable solvate thereof, to a human or non-human mammal in need thereof.
  • the active ingredient may be administered as a pharmaceutical composition hereinbefore defined, and this forms a particular aspect of the present invention.
  • the compound of formula G In treating hyperglycaemic or obese humans the compound of formula G). or a pharmaceutically acceptable salt, ester or amide thereof; or a pharmaceutically acceptable solvate thereof, may be taken in doses, such as those described above, one to six times a day in a manner such mat the total daily dose for a 70 kg adult will generally be in the range of from 0.1 to 6000 mg, and more usually about 1 to 1500 mg.
  • the treatment regimens for atherosclerosis are generally as described for hyperglycaemia. .
  • die active ingredient may be adminstered by mouth, usually once or twice a day and in an amount in the range of from about 0.025 mg/kg to 25 mg/kg, for example 0.1 mg/kg to 20 mg kg.
  • the present invention also provides a method for increasing weight gain and/or improving the feed utilisation efficiency and/or increasing lean body mass and/or decreasing birth mortality rate and increasing post/natal survival rate; of livestock, which method comprises the administration to livestock of an effective non-toxic amount of a compound of formula (I) or a veterinarily acceptable salt, ester or amide thereof, or a veterinarily acceptable solvate thereof.
  • the compounds of formula (I) and the veterinarily acceptable salts thereof or a veterinarily acceptable solvate thereof may be administered to any livestock in the abovementioned method, ti ey are particularly suitable for increasing weight gain and/or feed utilisation efficiency and/or lean body mass and/or decreasing birth mortality rate and increasing post-natal survival rate; in poultry, especially turkeys and chickens, catde, pigs and sheep.
  • d e compounds of formula (I) or veterinarily acceptable salt, ester or amide thereofs thereof will normally be administered orally although non-oral modes of administration, for example injection or implantation, are also envisaged.
  • the compounds are administered in the feed-stuff or drinking water provided for the livestock.
  • these are administered in the feed-stuff at from 10- ppm - 500ppm of total daily fed intake, more usually O.Olppm to 250ppm, suitably less than lOOppm.
  • the particular formulations used will of course depend upon d e mode of administration but will be those used conventionally in die mode of administration chosen.
  • the present invention also provides a veterinarily acceptable premix formulation comprising a compound of formula (I), or a veterinarily acceptable salt, ester or amide tiiereof thereof; or a veterinarily acceptable solvate thereof, in association with a veterinarily acceptable carrier therefore.
  • Suitable earners are inert conventional agents such as powdered starch. Other conventional feed-stuff premix carriers may also be employed.
  • the tide compound was obtained from d ⁇ sopropyl 4-(2-oxopropyl)phenoxymethyl- phosphonate (3.0 g, 9.1 mMol), R- ⁇ -methylbenzylamine (1.1 g, 9.1 mMol) and platinum IV oxide (25 mg) in ethanol (30 ml) by an analogous procedure to that described in Procedure 4 after chromatography over silica gel eluting with dichlorometiiane containing 3% methanol. !H NMR ⁇ (CDCI3)
  • the tide compound was obtained from di-n-butyl 4-(2-oxopropyl)phenoxymethyl- phosphonate (4.0 g, 10.5 mMol), R- ⁇ -methylbenzylamine (1.3 g, 10.5 mMol) and platinum IV oxide (30 mg) in ethanol (40 ml) by an analogous procedure to that described in Procedure 4 after chromatography over silica gel eluting with dichlorometiiane containing 3% methanol.
  • the tide compound was obtained from (RR)-di-n-butyl 4-[2-[2-(l-phenylethyl)- amino]propyl]phenoxymethylphosphonate (0.8 g, 1.8 mMol), 10% palladium on charcoal (300 mg) and ammonium formate (0.5 g, 7.9 mMol) in methanol (30 ml) by an analogous procedure to tiiat described in Procedure 5.
  • the tide compound was obtained from diisopropylphosphite (12.9 ml, 77.5 mmol), 4-fluorobenzaldehyde (10 ml, 77.4 mMol) paraformaldehyde (2.4 g, 77.4 mMol) and sodium hydride (3.29 g of a 60% dispersion in oil) by analogous procedure to that described in Procedure 1. Chromatography over silica gel eluting wid dichloromethane containing 3% methanol yielded a pale yellow oil. *H NMR ⁇ (CDCI3)
  • the title compound was obtained from di-n-butylphosphite (15.1 ml, 77.4 mMol), 4- fluorobenzaldehyde (2.4 g, 77.4 Mol), paraformaldehyde (2.4 g, 77.4 mMol) and sodium hydride (3.29 g of a 60% dispersion in oil) by an analogous procedure to that described in Procedure 1. Chromatography over silica gel eluting with dichloromethane containing 3% methanol yielded a pale yellow oil.
  • the tide compound was obtained in a two step sequence.
  • Initial treatment of di-n- butyl 4-formylphenoxymethylphosphonate (11.39 g, 32.4 mMol), n-butylamine (10 ml, 100 mMol) and benzene (70 ml) gave the N-butylimine.
  • the intermediate thus generated was tiien reacted widi nitroethane (10 ml) and acetic acid (20 ml) by an analogous procedure to tiiat described in Procedure 2.
  • the title compound was obtained from di-n-butyl 4-(2-nitropropenyl)phenoxymethyl- phosphonate (12.1 g, 30 mMol), iron powder (15 g), methanol (70 ml), water (20 ml) and acetic acid (50 ml) by an analogous procedure to diat described in Procedure 3. Chromatography over silica gel eluting wid hexane containing 30% etiiyl acetate gave a pale yellow oil.
  • the tide compound was obtained from (RR)-di-n-butyl 4-[2-[2-(3-chlorophenyl)-2- hydroxyethylamino]propyl]phenoxymethylphosphonate (240 mg, 0.46 mMol), 2M aqueous sodium hydroxide (2 ml) and dioxane (2 ml) by an analogous procedure to tiiat described in Example 2. After chromatography over Cjg reverse phase silica gel eluting with water containing 20% isopropanol a white powder was obtained, mp >300°C. -H NMR ⁇ (CD 3 OD)
  • the tide compound was obtained from (R) triethyl 4-[2-aminopropyl]- phenoxyphosphonoacetate (0.56 g,1.5 mMol), N-trimethylsilylacetamide (200 mg,1.5 mMol), (R)-3-chlorophenyloxirane (230 mg,1.5 mMol) and dimethyl sulphoxide (4 ml) by an analogous procedure to tiiat described in Example 3. Chromatography over silica gel eluting with dichlorometiiane containing 3% methanol yielded a pale yellow oil. l U NMR ⁇ (CDCI3 D2O)
  • Rat white adipocytes were prepared by collagenase digestion as described by
  • Lipoiysis stimulated by ⁇ -agonists was measured over 30 minutes in medium where d e adenosine concentration was controlled; die medium included (-)-N ⁇ -(2- phenylisopropyl)adenosine (lOOnM) + deaminase as decribed by Honnor et al.
  • Rat uterine horns were removed and bisected longitudinally. Each tissue was tied to a glass tissue holder and placed in Krebs-Henseleit solution in a 30 ml organ bath as before. Tissues were placed under a resting tension of lg and allowed to equilibrate. Each uterine strip was pre-contracted by die addition of 40mM K + to the bath to produced a steady tonic contraction, ⁇ -agonists were then added to die badi in a cumulative manner and results were expressed as percentage inhibition of contraction.

Landscapes

  • Health & Medical Sciences (AREA)
  • Chemical & Material Sciences (AREA)
  • Organic Chemistry (AREA)
  • General Health & Medical Sciences (AREA)
  • Life Sciences & Earth Sciences (AREA)
  • Engineering & Computer Science (AREA)
  • Bioinformatics & Cheminformatics (AREA)
  • Chemical Kinetics & Catalysis (AREA)
  • Molecular Biology (AREA)
  • General Chemical & Material Sciences (AREA)
  • Medicinal Chemistry (AREA)
  • Nuclear Medicine, Radiotherapy & Molecular Imaging (AREA)
  • Biochemistry (AREA)
  • Pharmacology & Pharmacy (AREA)
  • Diabetes (AREA)
  • Animal Behavior & Ethology (AREA)
  • Veterinary Medicine (AREA)
  • Public Health (AREA)
  • Obesity (AREA)
  • Hematology (AREA)
  • Endocrinology (AREA)
  • Emergency Medicine (AREA)
  • Heart & Thoracic Surgery (AREA)
  • Child & Adolescent Psychology (AREA)
  • Cardiology (AREA)
  • Vascular Medicine (AREA)
  • Urology & Nephrology (AREA)
  • Reproductive Health (AREA)
  • Pharmaceuticals Containing Other Organic And Inorganic Compounds (AREA)
  • Acyclic And Carbocyclic Compounds In Medicinal Compositions (AREA)
  • Organic Low-Molecular-Weight Compounds And Preparation Thereof (AREA)
  • Medicines That Contain Protein Lipid Enzymes And Other Medicines (AREA)
  • Saccharide Compounds (AREA)
  • Hydrogenated Pyridines (AREA)

Abstract

A compound of formula (I) or a pharmaceutically acceptable salt, ester or amide thereof, or a pharmaceutically acceptable solvate thereof, wherein R0 represents a substituted or unsubstituted aryl group; R1 represents hydrogen or an alkyl group; R2 represents a moiety of formula (a) wherein R?4 and R5¿ each independently represent hydrogen, alkyl, hydroxyalkyl, cycloalkyl or R4 together with R5 represents (CH¿2?)n wherein n is 2, 3 or 4; T represents hydrogen, nitrile or a group -CO.R?s¿ wherein Rs represents hydroxy, alkoxy or a group -NRtRu wherein R?t and Ru¿ independently represent hydrogen, alkyl or R?t and Ru¿ together with the nitrogen to which they are attached represent a saturated heterocyclic group; and R3 represents hydrogen, halogen, alkyl or alkoxy; a process for the preparation of such a compound, a pharmaceutical composition comprising such a compound and the use of such a compound in medicine.

Description

Phosphonated aryl ethanol ami ne compounds wi th anti -hypergly- cemi c and /or anti -obesi ty acti vi ty
This invention relates to novel compounds, to a process for preparing such compounds, to pharmaceutical compositions containing such compounds and to the use of such compounds and compositions in medicine and agriculture.
European patent application, publication number 0455006 discloses certain arylethanolamine derivatives which are stated to possess anti-diabetic and/or anti- hyperglycaemic and/or anti-obesity activity. It has been discovered that a series of novel phosphonated arylethanolamine derivatives have good β-adrenoreceptor agonist activity and in particular show good selectivity for β3-adrenoreceptors versus βi- or β2-adrenoreceptors. These compounds arc therefore indicated to have good anti-hyperglycaemic and/or anti- obesity activity coupled with especially good selectivity from cardiac and tremorigenic side effects.
They are also indicated to have potential in the treatment of gastrointestinal diseases such as irritable bowel syndrome.
These compounds also have potential as growth promoters for livestock and for decreasing birth mortality rate and increasing the post-natal survival rate in livestock. These compounds may also be of use in increasing the high-density- lipoprotein (HDL) cholesterol concentration and decreasing the triglyceride concentration in human blood serum and are therefore of potential use in the treatment and/or prophylaxis of atherosclerosis. They are also indicated to be useful for the treatment of hyperinsulinaemia. Accordingly the present invention provides a compound of formula (I):
Figure imgf000003_0001
(I)
or a pharmaceutically acceptable salt, ester or amide thereof thereof, or a pharmaceutically acceptable solvate thereof, wherein,
R° represents a substituted or unsubstituted aryl group; R.1 represents hydrogen or an alkyl group; R- represents a moiety of formula (a):
Figure imgf000004_0001
wherein R^ and R^ each independently represent hydrogen, alkyl, hydroxyalkyl, cycloalkyl or R^ together with R^ represents (CH2)n wherein n is 2, 3 or 4; T represents hydrogen, nitrile or a group -CO.Rs wherein Rs represents hydroxy, alkoxy or a group -NRlRu wherein R and Ru independently represent hydrogen, alkyl or Rl and Ru together with the nitrogen to which they are attached represent a saturated heterocyclic group; and
R- represents hydrogen, halogen, alkyl or alkoxy.
Suitably, R° represents a phenyl group optionally substituted with halogen or a hydroxy, examples include chlorophenyl, especially 3-chlorophenyl, and hydroxyphenyl, especially 3,4-bis-hydroxyphenyl. Suitably, R* is an alkyl group.
When R! is alkyl, it is favourably a C^.g alkyl group, especially a methyl group.
Suitably, R^ and R-5 each independently represent alkyl, for example ethyl propyl, or butyl. Preferably, R^ is alkyl, for example ethyl, propyl or butyl, especially ethyl, and R- is hydrogen.
Suitably, R- represents hydrogen.
Suitably, T represents hydrogen or a group -CO.Rs wherein Rs is hydroxy or alkoxy. Preferably, T represents hydrogen.
The compounds of formula (I) have one or two asymmetric carbon atoms, marked with an asterisk (*) or two asterisks (**) in the formula. These compounds may therefore exist in up to four stereoisomeric forms. The present invention encompasses all stereoisomers of the compounds of the general formula (I) whether free from other isomers, or admixed with other isomers in any proportion, such as mixtures of diastereoisomers and racemic mixtures of enantiomers.
Preferably, the asymmetric carbon atom indicated by a single asterisk (*) is in the R-configuration.
Preferably, the asymmetric carbon atom indicated by two asterisks (**) is in the R-configuration. One suitable form of a compound of formula (I) is a diastereomeric mixture of the RR and SS enantiomers.
One preferred form of a compound of formula (I) is the RR enantiomer. The term 'alkyl' when used alone or when forming part of other groups (such as the 'alkoxy' group) includes straight- or branched-chain alkyl groups containing 1 to 12 carbon atoms, suitably 1 to 6 carbon atoms, examples include methyl, ethyl, n-propyl, isopropyl, n-butyl, isobutyl or tert-butyl group.
The term 'cycloalkyl' includes C3_ cycloalkyl groups, especially C5 or Cg cycloalkyl groups. When used herein the term "halogen" refers to fluorine, chlorine, bromine and iodine, preferably chlorine.
When used herein the term 'aryl' includes phenyl and naphthyl optionally substituted with up to five, preferably up to three, groups selected from halogen, alkyl, phenyl, alkoxy, haloalkyl, hydroxyalkyl, hydroxy, amino, nitro, carboxy and pharmaceutically acceptable salts, esters and amides thereof, alkoxycarbonylalkyl, alkylcarbonyloxy and alkylcarbonyl groups.
A preferred aryl group is a substituted or unsubstituted phenyl group. Preferred optional substituents for the aryl group include up to three substituents selected from halogen, hydroxy, alkoxy, hydroxyalkyl and amino. Suitable pharmaceutically acceptable esters of carboxyl groups include alkyl esters, especially Cι_6 alkyl esters such as methyl.
Suitable pharmaceutically acceptable amides are those of formula -CONRxRy wherein R" and RY each independently represent hydrogen, alkyl or alkoxyalkyl.
Suitable pharmaceutically acceptable salts include acid addition salts, salts of carboxy groups and salts of phosphonic acid groups.
Suitable pharmaceutically acceptable acid addition salts include salts with inorganic acids such, for example, as hydrochloric acid, hydrobromic acid, orthophosphoric acid or sulphuric acid, or with organic acids such, for example as methanesulphonic acid, toluenesulophonic acid, acetic acid, propionic acid, lactic acid, citric acid, fumaric acid, malic acid, succinic acid, salicylic acid, maleic acid or acetylsalicylic acid.
Suitable pharmaceutically acceptable salts of carboxy groups or phosphonic acid groups include metal salts, such as for example aluminium, alkali metal salts such as sodium or potassium, alkaline earth metal salts such as calcium or magnesium and ammonium or substituted ammonium salts, for example those with C g alkylamines such as triethylamine, hydroxy-Cι_6 alkylamines such as 2-hydroxyethylamine, bis-(2- hydroxyethyl)-amine or tri-(2-hydroxyethyl)-amine, cycloalkylamines such as bicyclohexylamine, or with procaine, dibenzylpiperidine, N-benzyl-β-phenethylamine, dehydroabietylamine, N,N'-bisdehydroabietylamine, glucamine, N-methylglucamine or bases of the pyridine type such as pyridine, collidine or quinoline.
Suitable pharmaceutically acceptable solvates are conventional solvates, preferably hydrates.
The invention also provides a process for the preparation of a compound of formula (I), or a pharmaceutically acceptable salt, ester or amide thereof or a pharmaceutically acceptable solvate thereof, which process comprises reducing a compound of formula (II):
Figure imgf000006_0001
(II)
wherein R°, Rl, R2 and R^ are as defined in relation to formula (I), and thereafter, if necessary, carrying out one or more of the following optional steps:
(i) converting one compound of formula (I) to another compound of formula (I); and (ii) preparing a pharmaceutically acceptable salt, ester or amide thereof of a compound of formula (I) or a pharmaceutically acceptable solvate thereof.
The reduction of the compound of formula (II) may be carried out using any suitable reduction procedure, for example by using catalytic reduction in the presence of hydrogen. Suitable catalysts include platinum oxide or 10% palladium on charcaol.
Suitable reduction conditions include using an alkanolic solvent such as methanol, at any temperature providing a convenient rate of formation of the required product, for example when using the platinum catalyst the reaction may conveniently be carried out at ambient temperature or when using the palladium catalyst the reaction may be carried out at a medium temperature such as 50°C, under a pressure of 1-5 atmospheres of hydrogen.
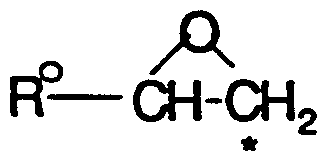
The compound of formula (II) may be prepared by reacting a compound of formula (III): OH o l
R -CH-CH2-NH2 *
(HI)
wherein R° is as defined in relation to formula (I) with a compound of formula (IV):
Figure imgf000007_0001
(IN)
wherein R2 and R~ are as defined in relation to formula (I). The reaction between compounds of formulae (III) and (IN) may be carried out under conventional animation conditions, for example in a solvent such as methanol or toluene.
Conveniently, the compound of formula (II) is prepared in-situ by reacting a compound of the above defined formula (III) with a compound of the above defined formula (IV) under reductive amination conditions.
Suitable reductive amination conditions include reacting the compounds of formula (III) and (IV) in an alkanolic solvent, such as methanol, in the presence of a suitable reduction catalyst.
Suitable reduction catalysts and reaction conditions are as described above for the reduction of the compound of formula (II).
A compound of formula (IV) may be prepared by reducing a compound of formula (V):
(V)
wherein R , R-- and R~ are as defined in relation to formula (I).
The reduction of the compound of formula (V) may conveniendy be carried out using iron powder in the presence of acetic acid in an aqueous solvent such as aqueous methanol, at any temperature providing a suitable rate of formation of the required product, generally at an elevated temperature and conveniently at the reflux temperature of the solvent.
A compound of formula (V) may be prepared by reacting a compound of formula (VI):
Figure imgf000008_0001
(ND
wherein, R-- and R3 are as defined in relation to formula (I), with nitroalkane. Generally, the carbon atom of the -CHO group of compound of formula (VI) is in an activated form, a suitable activated form being provided by forming an imine of the said carbonyl group: The imine may be prepared by reacting the compound of formula (VI) with an amine, suitably a primary alkyl amine such as n-butylamine. The reaction of the compound of formula (VI) and the amine may be carried out in any suitable solvent, such as toluene, at any temperature providing a suitable rate of formation of the required product, generally at an elevated temperature such as the reflux temperature of the solvent; and preferably in the presence of a catalytic amount of toluenesulphonic acid.
The reaction between the compound of formula (VI), and when it is in the form of an imine, and nitroalkane may be carried out in glacial acetic acid, preferably in the presence of an ammonium acetate catalyst, generally at an elevated temperature such as in the range of from 60°C to 120°C, for example 100°C.
A compound of formula (VI), wherein T is hydrogen, may be prepared from a compound of formula (Nil):
Figure imgf000008_0002
(Nil)
wherein R~ is as defined in relation to formula (I) and L* is a leaving group or atom, generally a fluorine atom, with an activated form of a compound of formula (VIII): HOCH2-P-OR4
5
OR
(VIII)
wherein R^ and R^ are as defined in relation to formula (I). A suitable activated form of a compound of formula (VIII) is an ionic form, such as a salted form, for example an alkali metal salted form.
An activated form of a compound of formula (VIII) may be prepared by use of the appropriate conventional procedure, for example a salted form may be prepared by treating the compound of formula (VHT) with a base such as an alkali metal hydride, for example sodium hydride.
The reaction between the compounds of formulae (VII) and (VIII) may be carried out in any suitable solvent, generally an aprotic solvent such as dimethylformamide or N-methylpyrrolidinone at a low to ambient temperature, for example in the range of from -15°C to 20°C, such as 5°C. A compound of formula (IV) may also be prepared by the reaction of a compound of formula (IX):
Figure imgf000009_0001
wherein R , R^ and R- are as defined in relation to formula (I) and J represents an oxo group or a protected oxo group, for example an ethan-l,2-dioxy group, with a compound of formula (X):
Figure imgf000009_0002
wherein R^, R^and T are as defined in relation to formula (I) in the presence of a rhodium salt , and thereafter, as required, removing any protecting group. A suitable rhodium salt is rhodium acetate.
The reaction between the compound of formula (IX) and the compound of formula (X) may be carried out under conventional carbene insertion conditions, in an inert solvent, such as benzene at any temperature providing a convenient rate of formation of the required product, generally at an elevated temperature, such as the reflux temperature of the solven; preferably the reaction is carried out under an inert atmosphere such as nitrogen. When J is a protected oxo group such as an ethan-l,2-dioxy group it may be removed by treatment with an aqueous acid, such as aqueous hydrochloric acid.
A compound of formula (I) wherein either the *CH2 or the **CH2 carbon is a chiral carbon may be prepared using conventional stereoselective chemistry from appropriate starting materialsis. One process for the preparation of a compound of formula (I) wherein both the *CH2 and the **CH2 carbon are chiral carbon atoms, or a pharmaceutically acceptable salt, ester or amide thereof or a pharmaceutically acceptable solvate thereof, comprises reacting a compound of formula (XI):
Figure imgf000010_0001
(XI) wherein R° is as defined in relation to formula (I), with a compound of formula (XII):
Figure imgf000010_0002
wherein Rl, R-^and R- are as defined in relation to formula (I); and thereafter, if necessary, carrying out one or more of the following optional steps:
(i) converting one compound of formula (I) to another compound of formula (I); and (ii) preparing a pharmaceutically acceptable salt, ester or amide thereof of a compound of formula (I) or a pharmaceutically acceptable solvate thereof.
Suitably, both the *CH2 and the **CH2 carbon atoms have the R stereochemistry.
Suitable reaction conditions for the reaction between the compounds of formulae (XI) and (XII) include using an aprotic solvent such as dimethylsulphoxide, at any temperature providing a convenient rate of formation of the required product, for example at an elevated temperature such as in the range of from 30°C to 100°C, for example 75°C: preferably the reaction is carried out under an inert atmosphere such as nitrogen. A compound of formula (XII) may be prepared by the hydrogenolysis of a compound of formula (XIII):
Figure imgf000011_0001
(XΠD
wherein R1, R-- and R- are as defined in relation to formula G) and the **CH carbon and ***CH carbon atoms are chiral carbon atoms.
Suitably, catalytic hydrogenolysis is used, using for example 10% palladium on charcaol in the presence of ammonium formate, suitably in an alkanolic solvent such as methanol, at any temperature providing a convenient rate of formation of the required product, for example at ambient temperature; preferably the reaction is carried out in an inert atmosphere, generably under nitrogen .
The compound of formula (XIII) may be prepared by stereoselective reduction of a compound of formula (XTV):
Figure imgf000011_0002
(XIV)
wherein R1, R^ and R-^ are as defined in relation to formula (I) and the ***CH carbon is a chiral carbon.
The reduction of the compound of formula (XTV) may be carried out using catalytic reduction in the presence of hydrogen. A preferred catalyst is platinum oxide.
Suitable reduction conditions include using an alkanol solvent such as methanol or ethanol, at any temperature providing a convenient rate of formation of the required product, conveniently at ambient temperature using a pressure of 1-5 atmospheres of hydrogen.
The compound of formula (XIV) may be prepared by reacting a compound of the above defined formula (IV) with R-α-methylbenzylamine.
The reaction between compounds of formulae (IV) and R-α- methylbenzylamine may be carried out under conventional amination conditions, for example those described above for the reaction between the compounds of formula (HI) and (TV). Conveniently, as for the above defined compound of formula (II), the compound of formula (XIV) is prepared inisiflL y reacting a compound of the above defined formula (TV) with R-α-methylbenzylamine under reductive amination conditions as described in relation to the compounds of formulae (III) and (TV).
Suitable conversions of one compound of formula (I) into another compound of formula (I) include converting one group OR4 into another group OR4 and/or converting one group OR^ into another group OR*.
Suitable conversions of one group OR4 into another group OR4 include:
(i) converting OR4 as hydroxy into OR4 as alkoxy; (ii) converting OR4 as alkoxy into OR4 as hydroxy; (iii) converting OR4 as alkoxy into OR4 as another alkoxy group.
The abovementioned conversion (i) may be carried out under conventional phosphonate alkylation methods, using for example the appropriate alcohol (R4OH) in the presence of hydrogen chloride.
The abovementioned conversion (ii) may be carried out using conventional phosphonate hydrolysis methods, for example by treating the appropriate compound of formula (I) with an alkaline metal hydroxide, such as sodium or lithium hydroxide. This reaction is especially useful when converting OR4 alkoxy into OR4 as hydroxy while leaving OR-5 as alkoxy. Alternatively, and especially when it is required to convert both OR4 and OR-5 from alkoxy into hydroxy, acidic hydrolysis may be used using for example hydrochloric acid.
The abovementioned conversion (iii) may be carried out by first converting OR4 as alkoxy into OR4 as hydroxy using the conditions set out in respect of the abovementioned conversion (ii), followed by converting the hydroxy group so formed into another alkoxy group, using the conditions set out in respect of the abovementioned conversion (i).
The abovementioned conversion (iii) is of particular use for preparing compounds of formula (I) wherein OR4 represents methoxy: such compounds are generally prepared from compounds of formula (I) wherein OR4 represents an alkyloxy group other than methoxy (suitably ethoxy) by first hydrolysing the relevant OR4 group (via conversion (ii)) to prepare a compound of formula (I) wherein OR4 represents hydroxy and thereafter methylating (via conversion (i)) to provide the required compound wherein OR4 represents methoxy.
Suitable conversions of one group OR^ into another group OR-^ include analogous conversions to those mentioned above in regard to converting one group OR4 into another group OR4. The protection of any reactive group or atom such as the variables R , R , R2 and R- , may be carried out at any appropriate stage in the aforementioned processes. Suitable protecting groups include those used conventionally in the art for the particular group or atom being protected. For example a keto group may be protected as an ketal. Protecting groups may be prepared and removed using die appropriate conventional procedure. For example a keto group may be protected as an ketal by reaction with an appropriate alcohol, for example ethan-l,2-diol, in the presence of an acid catalyst, such as 4-toluenesulphoπic acid. The said ketal may be removed by use of acidic hydrolysis, for example treatment with hydrochloric acid. A leaving group or atom is any group or atom that will, under the reaction conditions, cleave from the starting material, thus promoting reaction at a specified site. Suitable examples of such groups unless othewise specified are halogen atoms, mesyloxy groups and tosyloxy groups.
The compounds of formulae (ID) and (VIII) are known compounds or they may be prepared by processes analogous to those used to prepare known compounds, for example the compounds of formula (LIT) may be prepared according to methods disclosed in J.Org.Chem., 1974, 3_9_, 914; and the compounds of formula (VIII) may be prepared according to methods disclosed in Phosphorus and Sulphur, 1978, ≤, 455. The compounds of formula (IX) are known compounds or they may be prepared by methods analogous to those used to prepare known compounds, for example those methods disclosed in European application, publication number 0052963 and United Kingdom patent number 2084577.
The compounds of formulae (X) are either known commercially available compounds or they may be prepared from such compounds by conventional procedures for example those disclosed in the Chem. Berichte 1968, 101. 3734 and The Chemistry of Phosphorus, (J. Emsley and D. Hall), Published by Harper and Row, London, 1976. For example the compounds wherein T represents -CO.RlRu may be prepared from the commercially available compounds wherein T is carboxyl or an ethyl ester thereof, by conventional amination reactions. The compounds of formula (X) wherein T is a nitrile group are conveniently prepared from α- chloroacetonitrile and P(OR)3 wherein R is alkyl, suitably ethyl, and thereafter interconverting R into R4 and or R-5 as required.
The compounds of formula (XI) are known compounds or they may be prepared by methods analogous to those used to prepare known compounds, for example those methods disclosed in European application, publication number 0262785.
The salts, esters, amides and solvates of the compounds mentioned herein may be produced by methods conventional in the art: For example, salts may be prepared by treating a compound of formula (I) with the appropriate acid.
Esters of carboxylic acids may be prepared by conventional esterification procedures, for example alkyl esters may be prepared by treating the required carboxylic acid with the appropriate alkanol, generally under acidic conditions. Amides may be prepared using conventional amidation procedures, for example amides of formula CONRsRl may be prepared by treating the relevant carboxylic acid with an amine of formula HNRSR^ wherein Rs and R- are as defined above. Alternatively, a C\. alkyl ester such as a methyl ester of the acid may be treated with an amine of the above defined formula HNRsRl to provide the required amide.
Compounds of formula (I) and pharmaceutically acceptable salt, ester or amide thereofs thereof; or a pharmaceutically acceptable solvate thereof, produced by the above processes, may be recovered by conventional methods.
If required mixtures of isomers of the compounds of d e invention may be separated into individual stereoisomers by conventional means, for example by the use of an optically active acid as a resolving agent Suitable optically active acids which maybe used as resolving agents are described in Topics in Stereochemistry', Vol. 6, Wiley Interscience, 1971, Allinger, N.L. and Eliel, W.L. Eds.
Alternatively, any enantiomer of a compound of die invention may be obtained by stereospecific synthesis using optically pure starting materials of known configuration.
The absolute configuration of compounds may be determined by conventional X-ray crystallographic techniques.
As previously indicated, the compounds of the present invention have valuable pharmacological properties:
The present invention accordingly provides a compound of formula (I) or a pharmaceutically acceptable salt, ester or amide thereof thereof, or a pharmaceutically acceptable solvate thereof, for use as an active therapeutic substance.
In one aspect, the present invention provides a compound of formula G)» or a pharmaceutically acceptable salt, ester or amide U ereof thereof, or a pharmaceutically acceptable solvate thereof, for use in the treatment of hyperglycaemia in human or non-human animals.
The present invention further provides a compound of formula (I), or pharmaceutically acceptable salt, ester or amide thereof thereof, or a pharmaceutically acceptable solvate thereof, for use in the treatment of obesity in human or non-human animals.
A compound of formula (I), or a pharmaceutically acceptable salt, ester or amide thereof thereof, or a pharmaceutically acceptable solvate thereof, may be administered β£i £g or, preferably, as a pharmaceutical composition also comprising a pharmaceutically acceptable carrier.
Accordingly, the present invention also provides a pharmaceutical composition comprising a compound of formula (I), or a pharmaceutically acceptable salt, ester or amide thereof thereof, or a pharmaceutically acceptable solvate thereof, and a pharmaceutically acceptable carrier therefor.
As used herein the term "pharmaceutically acceptable" embraces compounds, compositions and ingredients for both human and veterinary use: for example the term "pharmaceutically acceptable salt" embraces a veterinarily acceptable salt The composition may, if desired, be in the form of a pack accompanied by written or printed instructions for use.
Usually the pharmaceutical compositions of the present invention will be adapted for oral administration, although compositions for administration by other routes, such as by injection, are also envisaged. Particularly suitable compositions for oral administration are unit dosage forms such as tablets and capsules. Other fixed unit dosage forms, such as powders presented in sachets, may also be used.
In accordance with conventional pharmaceutical practice the carrier may comprise a diluent, filler, disintegrant, wetting agent, lubricant, colourant, flavourant or other conventional adjuvant.
Typical carriers include, for example, microcrystalline cellulose, starch, sodium starch glycollate, polyvinylpyrrolidone, polyvinylpolypyrrolidone, magnesium stearate or sodium lauryl sulphate.
Most suitably the composition will be formulated in unit dose form. Such unit dose will normally contain an amount of the active ingredient in d e range of from 0.1 to 1000 mg, more usually 2-100 mg or 0.1 to 500 mg, and more especially 0.1 to 250 mg.
The present invention further provides a method for treating hyperglycaemia in a human or non-human mammal, which comprises administering an effective, non-toxic, amount of a compound of formula (I), or a pharmaceutically acceptable salt, ester or amide thereof thereof, or a pharmaceutically acceptable solvate thereof, to a hyperglycaemic human or non-human mammal in need tiiereof.
The present invention further provides a method for treating obesity or for the treatment and or prophylaxis of atherosclerosis in a human or non-human mammal, which comprises administering an effective, non-toxic, amount of a compound of formula (I), or a pharmaceutically acceptable salt, ester or amide thereof thereof, or a pharmaceutically acceptable solvate thereof, to a human or non-human mammal in need thereof.
Conveniently, the active ingredient may be administered as a pharmaceutical composition hereinbefore defined, and this forms a particular aspect of the present invention.
In treating hyperglycaemic or obese humans the compound of formula G). or a pharmaceutically acceptable salt, ester or amide thereof; or a pharmaceutically acceptable solvate thereof, may be taken in doses, such as those described above, one to six times a day in a manner such mat the total daily dose for a 70 kg adult will generally be in the range of from 0.1 to 6000 mg, and more usually about 1 to 1500 mg. The treatment regimens for atherosclerosis are generally as described for hyperglycaemia. .
In treating non-human mammals, especially dogs, die active ingredient may be adminstered by mouth, usually once or twice a day and in an amount in the range of from about 0.025 mg/kg to 25 mg/kg, for example 0.1 mg/kg to 20 mg kg. In a further aspect the present invention also provides a method for increasing weight gain and/or improving the feed utilisation efficiency and/or increasing lean body mass and/or decreasing birth mortality rate and increasing post/natal survival rate; of livestock, which method comprises the administration to livestock of an effective non-toxic amount of a compound of formula (I) or a veterinarily acceptable salt, ester or amide thereof thereof, or a veterinarily acceptable solvate thereof. Whilst the compounds of formula (I) and the veterinarily acceptable salts thereof or a veterinarily acceptable solvate thereof, may be administered to any livestock in the abovementioned method, ti ey are particularly suitable for increasing weight gain and/or feed utilisation efficiency and/or lean body mass and/or decreasing birth mortality rate and increasing post-natal survival rate; in poultry, especially turkeys and chickens, catde, pigs and sheep.
In the preceding method d e compounds of formula (I) or veterinarily acceptable salt, ester or amide thereofs thereof will normally be administered orally although non-oral modes of administration, for example injection or implantation, are also envisaged. Suitably the compounds are administered in the feed-stuff or drinking water provided for the livestock. Conveniently these are administered in the feed-stuff at from 10- ppm - 500ppm of total daily fed intake, more usually O.Olppm to 250ppm, suitably less than lOOppm. The particular formulations used will of course depend upon d e mode of administration but will be those used conventionally in die mode of administration chosen. For administration in feed-stuff die drugs are conveniendy formulated as a premix in association with a suitable carrier. Accordingly, the present invention also provides a veterinarily acceptable premix formulation comprising a compound of formula (I), or a veterinarily acceptable salt, ester or amide tiiereof thereof; or a veterinarily acceptable solvate thereof, in association with a veterinarily acceptable carrier therefore.
Suitable earners are inert conventional agents such as powdered starch. Other conventional feed-stuff premix carriers may also be employed.
No unacceptable toxicological effects are expected when compounds of die invention are administered in accordance with the present invention.
The following Examples illustrate the invention but do not limit it in any way.
Procedure 1
Diethyl 4-formyIphenoxymethylphosphonate
Figure imgf000018_0001
To a suspension of sodium hydride (6.58g, of a 60% dispersion in oil, previously washed with hexane) in dimethylformamide (100ml) was added dropwise diediyl phosphite (21.44g, 0.155 Mol). The resultant mixture was stirred for 0.5h at room temperature then treated with paraformaldehyde (4.8g, O.I6M0I) in one portion. After stirring for 1.5h, the mixture was cooled to 5°C and 4-fluorobenzaldehyde (19.22g, 0.154Mol) was added dropwise. After 1.5h, the mixture was poured into water (500ml) saturated with ammonium chloride. The organic material was extracted with diediyl etiier (3x150ml), the combined organic phases were washed with water (3x100ml), dried and evaporated to yield a dark yellow oil.
Chromatography on silica gel eluting with 0-25% ethyl acetate in hexane to yield d e tide compound as a pale yellow oil.
*H - nmr δ (CDCI3)
9.87 (1H, s); 7.85 (2H, d, J=10Hz); 7.05 (2H, d, J=10Hz); 4.5-3.9 (6H, complex multiplet); 1.33 (6H, t, J=7.00Hz).
Procedure 2
Diethyl 4-(2-nitropropenyl)phenoxymethylphosphonate
Figure imgf000018_0002
A mixture of diethyl 4-formylphenoxymethylphosphonate (ll.Og, 42.2mMol), n- butylamine (3.1g, 42.4mMol) and 4-toluenesulphonic acid (50mg) dissolved in toluene (75ml) was heated under reflux for 1.5h with concomitant removal of water. The reaction mixture was cooled and die solvent evaporated to yield a brown oil (12.1g). This oil was dissolved in glacial acetic acid (40ml) containing nitroethane (3.1ml, 3.2g, 42.7mMol) and heated at 100° for 1.5h. After cooling, the reaction mixture was poured into water (250ml) and extracted with ethyl acetate (2x150ml). The combined organic extracts were washed with water (2x100ml), dried and evaporated to yield a dark yellow oil (13.2g). Chromatography on silica gel eluting with 0-2% methanol in dichloromethane in gave the tide compound as a light yellow oil. l - nmr δ (CDC13)
8.03 (1H, s); 7.42 (2H, d, J=10Hz); 7.06 (2H, d, J=10Hz); 4.5-3.9 (6H, complex multiplet); 2.38 (3H, s); 1.38 (6H, t, J=7.0Hz).
Procedure 3
Diethyl 4-(2-oxopropyl)phenoxvmethylphosphonate
Figure imgf000019_0001
To a mixture of diediyl 4-(2-nitropropenyl)phenoxymethylphosphonate (6.9g 21.0mMol) and iron powder (15g, 267mMol) in methanol (75ml) and water (25ml) under reflux was added acetic acid (50ml) dropwise. After 3h the reaction mixture was cooled and filtered through celite, the methanol was evaporated and die residue poured into water/ediyl acetate, filtered through celite and die organic phase separated. The aqueous phase was extracted once with ethyl acetate and die combined organic layers washed widi water (2x100ml), dried and evaporated to yield a dark red oil. Chromatography on silica gel eluting with 0-25% ethyl acetate in hexane gave the title compound as a light yellow oil. *H - nmr δ (CDCI3) 7.13 (2H, d, J=8.8Hz); 6.93 (2H, d, J=8.8Hz); 4.3-4.1 (6H, complex multiple!); 3.65 (2H, s); 2.15 (3H, s); 1.36 (6H, t, J=7.15Hz).
Procedure 4
(RR)-Diethyl 4-[2-[2-(l-phenyIethyl)amino]propyl]phenoxymethylphosphonate
Figure imgf000020_0001
A solution of diethyl 4-(2-oxopropyl)phenoxymedιylphosphonate (3.00 g, 10 mMol) and R- α -methylbenzylamine (1.21 g, 10 mMol) dissolved in methanol (50 ml) was hydrogenated in the presence of platinum IV oxide (25 mg) at atmospheric pressure and room temperature. After 16 h the mixture was filtered and d e solvent evaporated to yield d e crude compound as a light yellow oil. Chromatography over silica gel eluting wid dichloromethane containing 3 % methanol gave the tide compound as a colourless oil. lH NMR δ (CDCI3)
7.36-7.22 (5H,m); 7.01 (2H,dJ=6.60Hz); 6.86 (2H,d,J=6.59Hz); 4.29-4.20 (6H,m); 3.91 (lH,q =6.60Hz); 2.84-2.69 (2H,m); 2.49-2.41 (lH,m); 1.55 (lH,brs); 1.36 (6H,U=7.42Hz); 1.29 (3H,dJ=6.59Hz); 0.89 (3H,d,J=6.32Hz).
Procedure 5
(R)-Diethyl 4-[2-aminopropyI]phenoxymethylphosphonate
Figure imgf000020_0002
A mixture of (RR)-diethyl 4-[2-[2-(l-phenylethyl)amino]propyl]phenoxymethyl- phosphonate (2.39 g, 5.9 mMol) ammonium formate (3 g, 47.5 mMol) and 10% palladium on charcoal (300 mg) in methanol (30 ml) was heated at reflux under an atmosphere of nitrogen. After 2 h the mixture was cooled and filtered, d e methanol evaporated and die residue partitioned between water (50 ml) and dichlorometiiane (50 ml). The aqueous layer was extracted once with dichloromethane (50 ml) and the combined organic layers dried and evaporated to yield the tide compound as colourless oil. *H NMR δ (CDC13)
7.12 (2H,dJ=8.53Hz); 6.90 (2H,d,J=8.52Hz); 4.29-4.18 (6H,m); 3.24-3.12 (lH,m); 2.75-2.45 (4H,m); 1.36 (6H, =7.15Hz); 1.13 (3H,dJ=6.32Hz).
Procedure 6
(RR)-DHsopropyl 4-[2-[2-(l-phenylethyl)amino]propyI]phenoxymethyl- phosphonate
Figure imgf000021_0001
The tide compound was obtained from dϋsopropyl 4-(2-oxopropyl)phenoxymethyl- phosphonate (3.0 g, 9.1 mMol), R-α-methylbenzylamine (1.1 g, 9.1 mMol) and platinum IV oxide (25 mg) in ethanol (30 ml) by an analogous procedure to that described in Procedure 4 after chromatography over silica gel eluting with dichlorometiiane containing 3% methanol. !H NMR δ (CDCI3)
7.35-7.21 (5H,m); 7.01 (2H,dJ=8.52Hz); 6.85 (2H,dJ=8.52Hz); 4.89-4.79 (2H,m); 4.19 (2H,dJ=10.17Hz); 3.91 (lH,q,J=6.6Hz); 2.81-2.70 (2H,m); 2.49-2.43 (lH,m); 1.51 (lH,brs); 1.35 (12H, =6.32Hz); 1.29 (3H,d,J=6.59Hz); 0.89 (3H,dJ=6.33Hz). Procedure 7
(R)-Diisopropyl 4-(2-aminopropyl)phenoxymethylphosphonate
Figure imgf000022_0001
The title compound was obtained from (RR)-diisopropyl 4-[2-[2-(l-phenylethyl)- amino] propyl] phenoxymethylphosphonate (2.48 g, 5.7 mMol), ammonium formate (2.5 g, 39 mMol) and 10% palladium charcoal (500 mg) by an analogous procedure to that described in Procedure 5. lH NMR δ (CDCI3)
7.10 (2H,dJ=8.53Hz); 6.89 (2H,d,J=8.80Hz); 4.89-4.77 (2H,m); 4.20 (2H,dJ=10.17Hz); 3.15-3.08 (lH,m); 2.65 (lH,dxd,J=5.5 and 13.47Hz); 2.45 (lH,dxd,J=8.25 and 13.48Hz); 1.46 (2H,brs); 1.36 (12H,t,J=6.32Hz); 1.10 (3H,dJ=6.05Hz).
Procedure 8
(RR)-Di-n-butyl 4-[2-[2-(l-phenylethyl)amino]propyI]phenoxymethyI- phosphonate
Figure imgf000022_0002
The tide compound was obtained from di-n-butyl 4-(2-oxopropyl)phenoxymethyl- phosphonate (4.0 g, 10.5 mMol), R-α-methylbenzylamine (1.3 g, 10.5 mMol) and platinum IV oxide (30 mg) in ethanol (40 ml) by an analogous procedure to that described in Procedure 4 after chromatography over silica gel eluting with dichlorometiiane containing 3% methanol. *H NMR δ (CDC13)
7.35-7.26 (5H,m); 7.01 (2H,dJ=8.52Hz); 6.85 (2H,( =8.79Hz); 4.25
(2H,(U= 10.17Hz); 4.19-4.05 (4H,m); 3.96-3.90 (lH,m); 2.81-2.70 (2H,m); 2.46-2.40 (lH,m); 1.70-1.62 (5H,m); 1.45-1.34 (4H,m); 1.29 (3H4J=6.87Hz); 0.92 (6H,U=7.42Hz); 0.89 (3H,d,J=6.35Hz).
Procedure 9
(R)-Di-n-butyl 4-(2-aminopropyl)phenoxymethylphosphonate
Figure imgf000023_0001
The tide compound was obtained from (RR)-di-n-butyl 4-[2-[2-(l-phenylethyl)- amino]propyl]phenoxymethylphosphonate (0.8 g, 1.8 mMol), 10% palladium on charcoal (300 mg) and ammonium formate (0.5 g, 7.9 mMol) in methanol (30 ml) by an analogous procedure to tiiat described in Procedure 5. *H NMR δ (CDCI3)
7.12 (2H,dJ=8.52Hz); 6.89 (2H,dJ=8.80Hz); 4.26 (2H,dJ=9.89Hz); 4.16 (4H,qJ=6.59Hz); 3.12 (lH,q =7.42Hz); 2.66 (lH,dxdJ=5.22 and 13.19Hz); 2.47 (lH,dxd =5.5 and 13.48Hz); 1.74-1.60 (6H,m); 1.48-1.40 (4H,m); 1.11 (3H,dJ=6.33Hz); 0.93 (6H,U=7.42Hz). Procedure 10
Diisopropyl 4-formylphenoxymethylphosphonate
Figure imgf000024_0001
The tide compound was obtained from diisopropylphosphite (12.9 ml, 77.5 mmol), 4-fluorobenzaldehyde (10 ml, 77.4 mMol) paraformaldehyde (2.4 g, 77.4 mMol) and sodium hydride (3.29 g of a 60% dispersion in oil) by analogous procedure to that described in Procedure 1. Chromatography over silica gel eluting wid dichloromethane containing 3% methanol yielded a pale yellow oil. *H NMR δ (CDCI3)
9.95 (lH,s); 7.99 (2H,dJ=8.72Hz); 7.22 (2H,d,J=8.79Hz); 5.05-4.92 (2H,m); 4.44 (2H,dJ=10.45Hz); 1.51 (12H, =6.05Hz).
Procedure 11
Di-n-butyl 4-formylphenoxymethyIphosphonate
Figure imgf000024_0002
The title compound was obtained from di-n-butylphosphite (15.1 ml, 77.4 mMol), 4- fluorobenzaldehyde (2.4 g, 77.4 Mol), paraformaldehyde (2.4 g, 77.4 mMol) and sodium hydride (3.29 g of a 60% dispersion in oil) by an analogous procedure to that described in Procedure 1. Chromatography over silica gel eluting with dichloromethane containing 3% methanol yielded a pale yellow oil. H NMR δ (CDCI3) 9.91 (lH,s); 7.84 (2H,dJ=8.52Hz); 7.08 (2H,dJ=8.8Hz); 4.36 (2H,dJ=10.45Hz); 4.17 (4H,qJ=6.6Hz); 1.74-1.63 (4H,m); 1.48-1.34 (4H,m); 0.93 (6H,U=7.43Hz).
Procedure 12
Diisopropyi 4-(2-nitropropenyl)phenoxymethylphosphonate
Figure imgf000025_0001
The title compound was obtained in a two step sequence. Initial treatment of diisopropyi 4-formylphenoxymethylphosphonate (6.7 g, 22.3 mMol) with n- butylamine (5 ml, 50 mMol) and benzene (70 ml) gave the N-butylimine. The intermediate thus generated was then reacted with nitroethane (10 ml) and acetic acid (20 ml) by an analogous procedure to that described in Procedure 2. iH NMR δ (CDCI3)
8.26 (lH,s); 7.11 (2H,dJ=8.5Hz); 6.89 (2H,dJ=8.5Hz); 4.95-4.70 (2H,m); 4.18 (2H,dJ=10.2Hz); 2.10 (3H,s); 1.32 (6H,dJ=7.5Hz).
Procedure 13
Di-n-butyl 4-(2-nitropropenyl)phenoxymethylphosphonate
Figure imgf000025_0002
The tide compound was obtained in a two step sequence. Initial treatment of di-n- butyl 4-formylphenoxymethylphosphonate (11.39 g, 32.4 mMol), n-butylamine (10 ml, 100 mMol) and benzene (70 ml) gave the N-butylimine. The intermediate thus generated was tiien reacted widi nitroethane (10 ml) and acetic acid (20 ml) by an analogous procedure to tiiat described in Procedure 2. *H NMR δ (CDCI3)
8.23 (lH,s); 7.60 (2H,dJ=8.5Hz); 7.31 (2H,d,J=8.7Hz); 4.5-4.1 (6H,m); 2.62 (3H,s,); 1.8-1.3 (8H,m); 1.11 (6H,t,J=7.5Hz).
Procedure 14
Diisopropyi 4-(2-oxopropyl)phenoxymethylphosphonate
Figure imgf000026_0001
The title compound was obtained from diisopropyi 4-[2-nitropropenyl)phenoxy- methylphosphonate (7.6 g, 21.5 mMol), iron powder (20 g), methanol (70 ml), water (20 ml) and acetic acid (30 ml) by an analogous procedure to that described in Procedure 3. Chromatography over silica gel eluting with hexane containing 30% ethyl acetate gave a pale yellow oil. iH NMR δ (CDCI3)
7.12 (2H,d,J=8.52Hz); 6.93 (2H,d,J=8.52Hz); 4.89-4.77 (2H,m); 4.20 (2H,dJ=10.45Hz); 3.64 (2H,s); 2.14 (3H,s); 1.36 (12H,t,J=6.04Hz).
Procedure 15
Di-n-butyl 4-(2-oxopropyl)phenoxymethylphosphonate
Figure imgf000027_0001
The title compound was obtained from di-n-butyl 4-(2-nitropropenyl)phenoxymethyl- phosphonate (12.1 g, 30 mMol), iron powder (15 g), methanol (70 ml), water (20 ml) and acetic acid (50 ml) by an analogous procedure to diat described in Procedure 3. Chromatography over silica gel eluting wid hexane containing 30% etiiyl acetate gave a pale yellow oil.
-H NMR δ (CDCI3)
7.13 (2H,d,J=8.53Hz); 6.92 (2H,d,J=8.8Hz); 4.27 (2H,d J=10.17Hz); 4.21-4.12 (4H,m); 3.64 (2H,s); 2.15 (3H,s); 1.74-1.63 (4H,m); 1.48-1.37 (4H,m); 0.93 (6H,t,J=7.42Hz).
Procedure 16
(R)-Triethyl 4-[2-aminopropyl]phenoxyphosphonoacetate
Figure imgf000027_0002
The title compound was obtained from (RR)-triethyl 4-[2-[2-(l- phenylethyl)amino]propyl]phenoxyphosphonoacetate (0.85 g,1.78 mMol), 10% palladium on charcoal (500 mg) and ammonium formate (2 g) in methanol (20 ml) by an analogous procedure to that described in Procedure 5. iH NMR δ (CDCI3)
7.10 (2H,d,J=8.52 Hz); 6.86 (2H,d,J=8.79 Hz); 5.00 (lH,d,J=18.97 Hz); 4.35-4.24 (6H,m); 3.23-3.05 (lH,m); 2.71-2.65 (lH,m); 2.52-2.39 (lH,m); 1.74-1.38 (2H,br m); 1.37 (6H,t,J=7.15 Hz); 1.29 (3H,U=7.14 Hz); 1.10 (3H,d,J=6.33 Hz).
Procedure 17
(RR)-Triethyl 4-[2-[2-(l-phenylethyl)amino]propyl]phenoxyphosphonoacetate
Figure imgf000028_0001
The title compound was obtained from triethyl 4-(2-oxopropyl)- phenoxyphosphonoacetate (1.5 g,4.0 mMol), (R)-α-methylbenzylamine (0.48 g,4.0 mMol) and platinum IV oxide (20 mg) in ethanol (25 ml) by an analogous procedure to that described in Procedure 4 after chromatography over silica gel eluting with dichloromethane containing 3% methanol. *H NMR δ (CDCI3)
7.33-7.23 (5H,m); 6.99 (2H,dJ=8.8 Hz); 6.82 (2H,dJ=8.52 Hz); 4.99 (lH,d,J=18.97 Hz); 4.34-4.24 (6H,m); 3.90 (lH,qJ=6.87 Hz); 2.77-2.63 (2H,m); 2.47-2.40 (lH,m); 1.75-1.50 (lH,br m); 1.37 (6H,tJ=7.15 Hz); 1.29 (3H,d,J=6.59 Hz); 1.28 (3H, =7.15 Hz); 0.89 (3H,dJ=6.33 Hz).
Procedure 18
Triethyl 4-(2-oxopropyl)phenoxyphosphonoacetate
Figure imgf000029_0001
To a solution of triethyl 4-[2-dioxalano-2-propyl]phenoxy]phosphonoacetate (2.27 g,5.5 mMol) in acetone (20 ml) was added 2 M aqueous hydrochloric acid (10 ml). After stirring at room temperature overnight the acetone was evaporated and die organic material extracted widi diethyl etiier (2 x 50 ml). The combined organic phases were dried and evaporated to yield the tide compound as a yellow oil.
-H NMR δ (CDC13)
7.26 (2H,d,J=8.8 Hz); 7.02 (2H,d,J=8.8 Hz); 5.15 (lH,dJ=18.97 Hz); 4.46-4.33 (6H,m); 3.77 (2H,s); 2.27 (3H,s); 1.51 (6H,t,J=6.87 Hz); 1.42 (3H,t,J=7.15 Hz).
Procedure 19
Triethyl 4-[2-[dioxaIano-2-propyl]phenoxyphosphonoacetate
Et),
Figure imgf000029_0002
To a solution of 2-(4-hydroxyphenylmethyl)-2-methyldioxalane (2.65 g, 13.66 mMol) in benzene (50 ml) was added rhodium II acetate dimer (60 mg,l mole %.). The solution was degassed under vacuum and then purged with nitrogen. The resultant green solution was heated to reflux and triethyl diazo phosphonoacetate (3.75 g,15.0 mMol) in benzene (10 ml) was added over a 1 h period. After 12 h die mixture was cooled and filtered, die benzene was evaporated to yield a dark yellow oil. Chromatography over silica gel eluting widi hexane containing 35% diediyl edier gave the title compound as a pale green oil. -H NMR δ (CDCI3)
7.26 (2H,dJ=8.8 Hz); 6.91 (2H,d,J=8.8 Hz); 5.07 (1H,<U=18.97 Hz); 4.42-4.25 (6H,m); 3.98-3.89 (2H,m); 3.85-3.75 (2H,m); 2.95 (2H,s); 2.25 (3H,s,); 1.44 (6H, =7.15 Hz); 1.35 (3H,tJ=7.15 Hz).
Procedure 20
Ethyl 4-(2-oxopropyl)phenoxymethylphosphonic acid
Figure imgf000030_0001
To suspension of ethyl 4-(2-dioxalano-2-propyl)phenoxymethylphosphonic acid (4.19 g,13.0 mMol) in acetone (25 ml) was added 1 M hydrochloric acid (10 ml). After stirring overnight the solvent was evaporated and die residue dissolved in a mixture of ethyl acetate and isopropanol (4:1). The solution was washed with brine, dried and evaporated to die product as a yellow oil.
-H NMR δ (D6 DMSO/D2O)
7.11 (2H,dJ=8.52 Hz); 6.94 (2H,dJ=8.8 Hz); 4.19 (2H,dJf=9.89 Hz); 4.16-4.02 (2H,m); 3.67 (2H,s); 2.10 (3H,s); 1.26 (3H,t,J=6.88 Hz). Procedure 21
Ethyl 4-[2-dioxalano-2-propyl]phenoxymethylphosphonic acid
Figure imgf000031_0001
A solution of diediyl 4-[2-dioxalano-2-propyl]phenoxymethylphosphonate (5 g,14.5 mMol) in dioxane (25 ml) was treated with 1 M lithium hydroxide (20 ml). After stirring overnight the dioxane was evaporated and the aqueous residue adjusted to pH 7 by addition of 2 M hydrochloric acid. Evaporation and drying under vaccum yielded d e tide compound as a white solid contaminated wid lid ium chloride. *H NMR δ (D6 DMSO)
7.19 (2H,dJ=8.52 Hz); 6.90 (2H,dJ=8.52 Hz); 4.0-3.8 (8H,m); 2.84 (2H,s); 1.24 (3H,s); 1.19 (3H,t,J=6.88 Hz).
Procedure 22
Diethyl 4-[2-dioxalano-2-propyl]phenoxymethylphosphonate
Figure imgf000031_0002
A mixture of diediyl 4-(2-oxopropyl)phenoxymed ylphosphonate (10 g,33.3 mMol), 1,2-ethanediol (1.84 ml,33.3 mMol) and 4-toluenesulphonic acid monohydrate (0.5 g) in benzene (75 ml) was heated under reflux widi removal of water. After 12 h a further equivalent of 1,2-ethanediol was added and heating continued for 10 h. The reaction mixture was cooled and washed with sodium bicarbonate solution. The organic layer was washed with water, dried and evaporated to yield the title compound as straw coloured oil. *H NMR δ (CDCI3)
7.20 (2H,dJ=8.53 Hz); 6.88 (2H,dJ=8.8 Hz); 4.37-4.18 (6H,m); 3.93-3.84 (2H,m); 3.80-3.71 (2H,m); 2.86 (2H,s); 1.37 (6H,t,J=7.14 Hz); 1.29 (3H,s).
Example 1
(RR,SS)-Diethyl 4-[2-[2-(3-chlorophenyl)-2-hydroxyethyl- amino]propyl]phenoxymethylphosphonate hydrochloride
Figure imgf000033_0001
A mixture of l-(3-chlorophenyl)-2-aminoethanol (0.86g, 5mMol) and diediyl 4-(2- oxopropyl)phenoxymethylphosphonate (1.5g, 5mMol) dissolved in medianol (25ml) was treated with platinum IV oxide (50mg) and stirred under an atmosphere of hydrogen for 12h. The mixture was filtered d rough celite and die medianol evaporated. The crude product was chromatographed on silica gel eluting with 0-2% medianol in dichlorometiiane to yield d e secondary amine as a viscous oil. This was dissolved in etiiyl acetate and treated with ethereal hydrogen chloride, cooled to - 15°C and fractional crystallisation gave d e product as the (RR,SS) diastereomer as a white solid, after filtering and drying under vacuum (m.p. 84-86°C ). lH - nmr δ (D6-DMSO)
9.3-9.0 (IH, br s, exchanges with D2O); 8.8-8.5 (IH, br s, exchanges widi D2O); 7.5- 7.3 (4H, m); 7.17 (2H, d, J=8.52Hz); 7.00 (2H, d, J=8.52Hz); 6.63 (IH, d, J=3.77Hz; exchanges with D2O); 4.39 (2H, d, J=9.62); 4.2-4.0 (4H, m); 3.5-3.0 (4H, complex multiplet); 2.61 (IH, t, J=10.45Hz); 1.25 (6H, t, J=7.15Hz); 1.10 (3H, d, J=6.32Hz).
Example 2
(RR,SS)-Sodium ethyl 4-[2-[2-(3-chlorophenyl)-2- hydroxyethylamino]propyl]phenoxymethylphosphonate
Figure imgf000034_0001
(RR,SS)-Diethyl 4-[2-[2-(3-chlorophenyl)-2-hydroxyetiιylamino]propyl]phenoxy- methylphosphonate hydrochloride (106mg, 0.22mMol) was suspended in dioxane (2ml) and treated with 2.5M aqueous sodium hydroxide solution (2.0ml). After stirring at room temperature for 24h the dioxane was evaporated and die residual solution adjusted to pH9. Chromatography on C18 reverse phase silica eluting with water then 25 - 50% isopropanol in water, gave the title compound as a white solid, m.p. 122-5°C. lH - nmr δ (CD3OD)
7.45 (IH, s); 7.4-7.3 (3H, m); 7.13 (2H, d, J=8.6Hz); 6.93 (2H, d, J=8.7Hz); 4.90 (IH, dxt, J=9.0Hz); 4.1-4.0 (4H, ml; 3.3-3.2 (IH, m); 3.2-2.9 (3H,m); 2.68 (IH, dxd, J=13.4Hz); 1.25 (3H, t, J=7.1Hz); 1.19 (3H, d, J=6.5Hz).
Example 3
(RR)-Diethyl 4-[2-[2-(3-chlorophenyI)-2-hydroxyethylamino]phenoxy- ethy I phosphonate
Figure imgf000034_0002
A mixture of (R)-diedιyl-4-(2-aminopropyl)phenoxymethylphosphonate (388 mg, 1.29 mMol) and trimethylsilyl acetamide (168 mg, 1.29 mMol) in dimediyl sulfoxide (2.5 ml) was stirred at room temperature. After 0.5 h d e mixture was treated wid (R)-3-chlorophenyloxirane (200 mg, 1.29 mMol) and heated at 75°C under nitrogen for 50 h. The resultant solution was cooled and partitioned between water (10 ml) and ethyl acetate (20 ml). After drying and evaporation die organic layer gave a brown oil, chromatography of which, over silica gel eluting with dichloromethane containing 4% methanol, gave the tide compound as a colourless oil. *H NMR δ (CDCI3)
7.34 (lH,s); 7.26-7.17 (3H,m); 7.09 (2H J=8.52Hz); 6.89 (2H,d,J=8.52Hz); 4.55 (lH,dxdJ=3.85 and 8.80 Hz); 4.30-4.19 (6H,m); 2.95-2.79 (2H,m); 2.75-2.57 (5H,m); 1.36 (6H,U=7.15Hz); 1.06 (3H,dJ=6.05Hz).
Example 4
(RR)-Ethyl sodium 4-[2-[2-(3-chlorophenyl)-2-hydroxyethylamino]propyl]- phenoxymethylphosphonate
Figure imgf000035_0001
The title compound was obtained from (RR)-diethyl 4-[2-[2-(3-chlorophenyl)-2- hydroxyethylamino]propyl]phenoxymethylphosphonate (110 mg, 0.24 mMol), 2 M aqueous sodium hydroxide (2 ml) and dioxane (4 ml) by an analogous procedure to that described in Example 2. After chromatography over Cjg reverse phase silica eluting with water isopropanol (3:1) a white powder was obtained, mp>290°C. -H NMR δ (CD3OD)
7.35-7.30 (3H,m); 7.26 (lH,s); 7.00 (2H,d,J=8.59Hz); 6.87 (2H,d,J=8.64Hz); 4.67 (lH, =6.83Hz); 4.15-4.00 (4H,m); 2.98 (lH,dxd,J=7.03 and 11.70Hz); 2.90-2.80 (lH,m); 2.70-2.60 (2H,m); 2.50 (lH,dxd =8.28 and 13.51Hz); 1.28 (3H,tJ=7.08Hz); 1.10 (3H,d,J=6.25Hz). Example 5
(RR)-Diisopropyl 4-[2-[2-(3-chlorophenyl)-2-hydroxyethylamino]propyl]- phenoxymethylphosphonate
Figure imgf000036_0001
The title compound was obtained from (R)-diisopropyl 4-[2-aminopropyl]- phenoxymediylphosphonate (600 mg, 1.82 mMol), N-trimethylsilylacetamide (267 mg, 1.87 mMol), (R)-3-chloroρhenyloxirane (290mg, 1.87mMol) and dimethyl sulphoxide (2 ml) by analogous procedure to tiiat described in Example 3. Chromatography over silica gel eluting widi dichlorometiiane containing 3% methanol yielded a colourless oil. lH NMR δ (CDCI3)
7.34 (lH,s); 7.26-7.21 (3H,m); 7.09 (2H,d,J=8.52Hz); 6.89 (2H,dJ=8.53Hz); 4.86- 4.76 (3H,m); 4.58 (lH,dxd-J=3.30 and 8.80Hz); 4.20 (2H,dJ=10.17Hz); 2.92-2.86 (2H,m); 2.70-2.62 (3H,m); 1.36 (12H,t,J=6.05Hz); 1.08 (3H,d,J=6.32Hz).
Example 6
(RR)-Isopropyl sodium 4-[2-[2-(3-chlorophenyl)-2-hydroxyethylamino]propyl]- phenoxymethylphosphonate
Figure imgf000036_0002
The title compound was obtained from (RR)-diisopropyl 4-[2-[2-(3-chlorophenyl)-2- hydroxyethylamino]propyl]phenoxymethylphosphonate (350 mg, 0.72 mMol), 2 M aqueous sodium hydroxide (2 ml) and dioxane (2 ml) by an analogous procedure to tiiat described Example 2. Chromatography over Cjg reverse phase silica eluting with water containing 30% isopropanol gave a white solid, mp 299-301°C. -H NMR δ (CD3OD)
7.35-7.26 (3H,m); 7.21-7.18 (lH,m); 7.03 (2H,J=8.6Hz); 6.88 (2HJ=8.6Hz); 4.30- 4.24 (lH,m); 4.13-4.06 (lH,m); 4.04 (2H,cU=10.1Hz); 2.95-2.80 (3H,m); 2.71-2.53 (2H,m); 1.26 (6H,tJ=6.2Hz); 1.08 (3H,cU=6.2Hz).
Example 7
(RR)-Di-n-butyl 4-[2-[2-(3-chloropheπyl)-2-hydroxyethylamino]propyI]- phenoxymethylphosphonate
Figure imgf000038_0001
The title compound was obtained from (R)-di-n-butyl 4-(2- aminopropyl)phenoxymethylphosphonate (560 mg, 1.64 mMol), N- trimethylsilylacetamide (240 mg, 1.82 mMol) and (R)-3-chlorophenyloxirane
(262 mg, 1.69 mMol) in dimediyl sulfoxide (5 ml) by an analogous procedure to that described in Example 3. Chromatography over silica gel eluting with dichlorometiiane containing 3% methanol yielded a pale yellow oil. -H NMR δ (CDCI3)
7.34 (lH,s); 7.26-7.20 (3H,m); 7.08 (2H,d,J=8.52Hz); 6.89 (2H,d,J=8.52Hz); 4.59- 4.51 (2H,m); 4.26 (2H,dJ=11.54Hz); 4.19-4.12 (4H,m); 2.90-2.85 (lH,m); 2.65-2.61 (2H,m); 2.25-2.00 (2H,brm); 1.71-1.63 (3H,m); 1.46-1.37 (4H,m); 1.08 (3H,dJ=6.32Hz); 0.93 (6H,t =7.42Hz).
Example 8
(RR)-n-Butyl sodium 4-[2-[2-(3-chloropheπyl)-2-hydroxyethylamino]propyI]- phenoxymethylphosphonate
Figure imgf000039_0001
The tide compound was obtained from (RR)-di-n-butyl 4-[2-[2-(3-chlorophenyl)-2- hydroxyethylamino]propyl]phenoxymethylphosphonate (240 mg, 0.46 mMol), 2M aqueous sodium hydroxide (2 ml) and dioxane (2 ml) by an analogous procedure to tiiat described in Example 2. After chromatography over Cjg reverse phase silica gel eluting with water containing 20% isopropanol a white powder was obtained, mp >300°C. -H NMR δ (CD3OD)
7.32-7.25 (3H,m); 7.16-7.13 (lH,m); 6.97 (2H,dJ=8.63Hz); 6.85 (2H,dJ=8.67Hz); 4.65 (lH, =6.85Hz); 4.08 (2H,dJ=9.86Hz); 4.01-3.94 (2H,m); 2.98 (lH,dxd,J=6.72 and 11.53Hz); 2.88-2.70 (lH,m); 2.67-2.52 (lH,m); 2.45 (2H,dxd,J=8.52 and 13.59Hz); 1.64-1.55 (2H,m); 1.43-1.33 (2H,m); 1.08 (3H,dJ=6.12Hz); 0.89 (3H,t,J=7.32Hz).
Example 9
(RR)-Disodium 4-[2-[2-(3-Chlorophenyl)-2-hydroxyethylamino]propyl]- phenoxymethylphosphonate
Figure imgf000039_0002
A mixture (RR) of diisopropyi 4-[2-[2-(3-chlorophenyl)-2-hydroxyethylaminopropyl- methylphosphonate (100 mg, 0.2 mMol) and 6 M aqueous hydrochloric acid (2.5 ml) was heated at 100°C under an atmosphere of nitrogen for 12h. After cooling and evaporation of die solvent the resultant green gum was treated widi aqueous saturated sodium bicarbonate (1 ml) and die residue chromatographed over C\~ reverse phase silica to yield die title compound as a white crytstalline powder, mp>300°C. - NMR δ D2O DCI
7.36-7.31 (3H,m); 7.27-7.21 (lH,m); 7.18 (2H,d,J=8.68Hz); 6.98 (2H,dJ=8.69Hz), 4.95 (lH,t,J=6.63Hz); 4.22 (2H,d,J=9.92Hz); 3.59-3.53 (lH,m); 3.32-3.13 (2H,m); 3.02-2.96 (lH,m); 2.84-2.78 (lH,m); 1.23 (3H,d,J=6.6Hz).
Example 10
(RR) Dilithium ethyl 4-[2-[2-(3-chlorophenyl)-2-hydroxyethylamino]propyl]- phenoxyphosphonoacetate
Figure imgf000040_0001
C02Li
The title compound was obtained from (RR)-triethyl 4-[2-[2-(3-chlorophenyl)-2- hydroxyethylamino]propyl]phenoxyphosphonoacetate (174 mg,0.37 mMol), 1 M lithium hydroxide (2 ml) and dioxane (3 ml) by an analogous procedure to that described in Example 2. After chromatography over Cjg reverse phase silica eluting with waterΛsopropanol (9:1) a white powder was obtained, mp>300°C. *H NMR δ (CD3OD/D2O)
7.43 (lH,s); 7.39-7.32 (2H,m); 7.32-7.29 (lH,m); 7.14 (2H,dxdJ=8.36 and 7.2 Hz); 6.92 (2H,dxd,J=8.36 and 7.2 Hz); 4.90-4.88 (lH,m); 4.68 (lH,d,J=18.1 Hz); 4.08- 4.01 (2H,m); 3.20-3.00 (2H,m); 2.95-2.80 (lH,m); 2.80-2.70 (lH,m); 1.24 (3H, =5.8 Hz); 1.18 (3H,d,J=12 Hz). Example 11
(RR) Triethyl 4-[2-[2-(3-chlorophenyl)-2-hydroxyethyIamino]propyl]- phenoxyphosphonoacetate
Figure imgf000041_0001
C02Et
The tide compound was obtained from (R) triethyl 4-[2-aminopropyl]- phenoxyphosphonoacetate (0.56 g,1.5 mMol), N-trimethylsilylacetamide (200 mg,1.5 mMol), (R)-3-chlorophenyloxirane (230 mg,1.5 mMol) and dimethyl sulphoxide (4 ml) by an analogous procedure to tiiat described in Example 3. Chromatography over silica gel eluting with dichlorometiiane containing 3% methanol yielded a pale yellow oil. lU NMR δ (CDCI3 D2O)
7.35 (lH,s); 7.3-7.1 (3H,m); 7.06 (2H,dJ=8.52 Hz); 6.86 (2H,dJ=8.52 Hz); 5.04
(lH,d,J=18.9 Hz); 4.65 (lH,m); 4.0-4.1 (6H,m); 3.0-2.8 (lH,m); 2.7-2.6 (3H,m); 2.0- 1.9 (lH,m); 1.37 (6H,U=7.15 Hz); 1.29 (3H,t,J=7.15 Hz); 1.08 (3H,dJ=6.32).
Example 12
(RR,RS) Ethyl 4-[2-[2-(3,4-dihydroxyphenyl)hydroxyethylamino]propyl]- phenoxymethylphosphonic acid
Figure imgf000041_0002
A mixture of ethyl 4-(2-oxopropyl)phenoxymethyl phosphonic acid (0.8 g,2.96 mMol) and (R)-noradrenaline (0.5 g,2.96 mMol) in methanol (50 ml) was stirred at room temperature for 0.5 h. Palladium on charcoal (10%,0.3 g) was men added and die resultant mixture hydrogenated at 50 psi, 50°C for 20 h. After filtration and evaporation of die solvent the crude product was chromatographed over Cj reverse phase silica eluting with water containing isopropranol 20% gave a pale yellow gum freeze drying of which yielded d e tide compound as a white fluffy solid, mp 170- 3°C. *H NMR δ (D2O)
7.28-7.10 (2H,m); 7.09-6.93 (2H,m); 6.91-6.81 (2H,m); 6.82-6.74 (lH,m); 4.18
(2H,dJ=9.5 Hz); 4.09-3.95 (2H,m); 3.60-3.48 (lH,m); 3.40-3.15 (2H,m); 3.00-2.80 (3H,m); 1.30 (3H,d,J=6.2 Hz); 1.26 (3H,t,J=6.8 Hz).
Pharmacological Data
Lipoiysis
Rat white adipocytes were prepared by collagenase digestion as described by
Rodbell (1964) with the modification of Honnor et al. (1985), in that adenosine (200nM) was included in die preparation to inhibit basal lipoiysis.
Lipoiysis stimulated by β-agonists was measured over 30 minutes in medium where d e adenosine concentration was controlled; die medium included (-)-N^-(2- phenylisopropyl)adenosine (lOOnM) + deaminase as decribed by Honnor et al.
(1985). The incubation was stopped by addition of trichloracetic acid (0.2ml of 10% w/v) to 1ml of cells. Glycerol was measured by die fluorimetric method of Boobis and Maugham (1983).
References
1. Rodbell M. (1964) J. Biol. Chem. 239, 375-380
2. Honnor R.C., Dhellon G.S. and Londos C. (1985) J. Biol. Chem. 260, 15122- 15129
3. Boobis L.H. and Maugham R.J. (1983) Clinica Chemica Acta 132, 173-179.
Phosphonic acid derivative of Example 2, EC50 = 77 nM (Intrinsic Activity = 0.95).
Agonist Activity at Rat βj and β Adrenoceptors In Vitro
βχ-Adrenoceptor Agonism
Female Sprague-Dawley rats (150-250g) were killed by a blow to die head and exsanguinated. Spontaneously beating right atria were removed by d e method of Broadley and Lumley (1977) and mounted on a glass tissue holder. Each tissue was placed in 30 ml organ baths at 37°C containing Kreb's-Henseleit solution. Each atrium was attached to an isometric transducer by cotton and placed under an initial resting tension of lg. Rate recordings from the spontaneous beating atria were obtained from the tension signal using a Lectromed Type 4522 ratemeter. All traces were recorded on a Lectromed M4 chart recorder, β-adrenoceptor agonists were tiien added to the Krebs medium in a cumulative fashion and d e results expressed as a percentage increase in atrial rate.
β2-Adrenoceptor Agonism
Rat uterine horns were removed and bisected longitudinally. Each tissue was tied to a glass tissue holder and placed in Krebs-Henseleit solution in a 30 ml organ bath as before. Tissues were placed under a resting tension of lg and allowed to equilibrate. Each uterine strip was pre-contracted by die addition of 40mM K+ to the bath to produced a steady tonic contraction, β-agonists were then added to die badi in a cumulative manner and results were expressed as percentage inhibition of contraction.
Agonist EC50 (atria) and IC50 (uteri) were calculated as the concentration of agonist producing 50% of their maximum increase in atrial rate or uterine relaxation. Relative intrinsic activity expressed as die maximal responses to test agonists relative to isoprenaline (=1.0) in both atria and uteri.
References
K . Broadley & P. Lumley (1977) Br. J. Pharmacol.52.51
Ki(nM)
Pi β2
Example 2 37580 10000

Claims

Claims
1. A compound of formula (I):
Figure imgf000045_0001
(I)
or a pharmaceutically acceptable salt, ester or amide thereof thereof, or a pharmaceutically acceptable solvate thereof, wherein, R° represents a substituted or unsubstituted aryl group; R! represents hydrogen or an alkyl group; R-- represents a moiety of formula (a):
O -CH-P-OR
I I 5
T OR
wherein R4 and R-5 each independentiy represent hydrogen, alkyl, hydroxyalkyl, cycloalkyl or R4 together with R represents (CH2)n wherein n is 2, 3 or 4; T represents hydrogen, nitrile or a group -CO.Rs wherein Rs represents hydroxy, alkoxy or a group -NRlRu wherein Rl and R~ independentiy represent hydrogen, alkyl or R* and Ru together with the nitrogen to which they are attached represent a saturated heterocyclic group; and R3 represents hydrogen, halogen, alkyl or alkoxy.
2. A compound according to claim 1, wherein R° represents a phenyl group optionally substituted with halogen or a hydroxy group.
3. A compound according to claim 1 or claim 2, wherein R is a methyl group.
4. A compound according to any one of claims 1 to 3, wherein R4 is alkyl and R5 is hydrogen.
5. A compound according to any one of claims 1 to 4, wherein T represents hydrogen.
6. A compound according to any one of claims 1 to 5, wherein Rx represents hydrogen.
7. A compound according to claim 1 as described in examples 1 to 12, or a pharmaceutically acceptable salt, ester or amide thereof or a pharmaceutically acceptable solvate thereof.
8. A process for the preparation of a compound of formula (I), or a pharmaceutically acceptable salt, ester or amide thereof or a pharmaceutically acceptable solvate thereof, which process comprises:
a) reducing a compound of formula GI):
Figure imgf000046_0001
wherein R°, R1, R2 and R3 are as defined in relation to formula (I); or
b) for the preparation of a compound of formula (I) wherein both the *CH2 and the **CH2 carbon are chiral carbon atoms, or a pharmaceutically acceptable salt, ester or amide thereof or a pharmaceutically acceptable solvate thereof, reacting a compound of formula (XI):
Figure imgf000046_0002
(XI) wherein R° is as defined in relation to formula (I), with a compound of formula (XII):
Figure imgf000047_0001
wherein R , R2and R~ are as defined in relation to formula G); and diereafter, if necessary, carrying out one or more of the following optional steps:
(i) converting one compound of formula (I) to another compound of formula (I); and (ii) preparing a pharmaceutically acceptable salt, ester or amide thereof of a compound of formula (I) or a pharmaceutically acceptable solvate thereof.
9. A pharmaceutical composition comprising a compound of formula (I), or a pharmaceutically acceptable salt, ester or amide thereof; or a pharmaceutically acceptable solvate thereof, and a pharmaceutically acceptable carrier therefor.
10. A compound of formula (I), or a pharmaceutically acceptable salt, ester or amide d ereof; or a pharmaceutically acceptable solvate thereof, for use as an active dierapeutic substance.
11. A compound of formula (I), or a pharmaceutically acceptable salt, ester or amide diereof; or a pharmaceutically acceptable solvate thereof, for use in the treatment of hyperglycaemia, obesity, atherosclerosis, hyperinsulinaemia and/or irritable bowel syndrome.
12. The use of a compound of formula G). or a pharmaceutically acceptable salt, ester or amide diereof; or a pharmaceutically acceptable solvate thereof, for the manufacture of a medicament for the treatment of hyperglycaemia, obesity, atherosclerosis, hyperinsulinaemia and/or irritable bowel syndrome.
13. A method for treating hyperglycaemia, obesity, atherosclerosis, hyperinsulinaemia and/or irritable bowel syndrome in a human or non-human mammal, which comprises administering an effective, non-toxic, amount of a compound of formula (I), or a pharmaceutically acceptable salt, ester or amide thereof; or a pharmaceutically acceptable solvate thereof, to the human or non-human mammal in need diereof.
14. A method for increasing weight gain and/or improving die feed utilisation efficiency and/or increasing lean body mass and or decreasing birth mortality rate and increasing post/natal survival rate; of livestock, which method comprises the administration to livestock of an effective non-toxic amount of a compound of formula (I) or a veterinarily acceptable salt, ester or amide thereof; or a veterinarily acceptable solvate diereof.
15. A veterinarily acceptable premix formulation comprising a compound of formula ( , or a veterinarily acceptable salt, ester or a ide diereof; or a veterinarily acceptable solvate thereof, in association wid a veterinarily acceptable carrier therefore.
PCT/GB1993/001554 1992-07-25 1993-07-22 Phosphonated arylethanolamine compounds with anti-hyperglycemic and/or anti-obesity activity Ceased WO1994002493A1 (en)

Priority Applications (4)

Application Number Priority Date Filing Date Title
DE69311118T DE69311118T2 (en) 1992-07-25 1993-07-22 PHOSPHONO-ARYLETHANOLAMINE COMPOUNDS WITH ANTI-HYPERGLYCEMIC AND / OR ANTIOBESIC EFFECT
EP93917898A EP0652884B1 (en) 1992-07-25 1993-07-22 Phosphonated arylethanolamine compounds with anti-hyperglycemic and/or anti-obesity activity
JP6504304A JPH07509458A (en) 1992-07-25 1993-07-22 Phosphonated arylethanolamine compounds with antihyperglycemic and/or antiobesity activity
AU47151/93A AU4715193A (en) 1992-07-25 1993-07-22 Phosphonated arylethanolamine compounds with anti-hyperglycemic and/or anti-obesity activity

Applications Claiming Priority (2)

Application Number Priority Date Filing Date Title
GB9215844.3 1992-07-25
GB929215844A GB9215844D0 (en) 1992-07-25 1992-07-25 Novel compounds

Publications (1)

Publication Number Publication Date
WO1994002493A1 true WO1994002493A1 (en) 1994-02-03

Family

ID=10719297

Family Applications (1)

Application Number Title Priority Date Filing Date
PCT/GB1993/001554 Ceased WO1994002493A1 (en) 1992-07-25 1993-07-22 Phosphonated arylethanolamine compounds with anti-hyperglycemic and/or anti-obesity activity

Country Status (13)

Country Link
US (1) US5519006A (en)
EP (1) EP0652884B1 (en)
JP (1) JPH07509458A (en)
CN (1) CN1101349A (en)
AT (1) ATE153666T1 (en)
AU (1) AU4715193A (en)
CA (1) CA2140916A1 (en)
DE (1) DE69311118T2 (en)
GB (1) GB9215844D0 (en)
MX (1) MX9304467A (en)
TW (1) TW249804B (en)
WO (1) WO1994002493A1 (en)
ZA (1) ZA935340B (en)

Cited By (13)

* Cited by examiner, † Cited by third party
Publication number Priority date Publication date Assignee Title
WO1995004047A1 (en) * 1993-07-31 1995-02-09 Smithkline Beecham Plc 2-benzoheterocyclyloxy or thiopropanolamine derivatives with adreno receptor agonist activity
WO1995007284A1 (en) * 1993-09-09 1995-03-16 Smithkline Beecham Plc Phosphinic acid derivatives with anti-hyper glycemic and/or anti-obesity activity
US5491134A (en) * 1994-09-16 1996-02-13 Bristol-Myers Squibb Company Sulfonic, phosphonic or phosphiniic acid β3 agonist derivatives
WO1996004233A1 (en) * 1994-07-29 1996-02-15 Smithkline Beecham Plc Aryloxy and arylthiopropanolamine derivatives useful as beta 3-adrenoreceptor agonists and antagonists of the beta 1 and beta 2-adrenoreceptors and pharmaceutical composition thereof
WO1996004234A1 (en) * 1994-07-29 1996-02-15 Smithkline Beecham Plc Aryloxy andarycthio propanolamine derivatives useful as b3-adrenoreceptor agonists and antagons of the b1 and b2 adrenoreceptors and pharmaceutical composition thereof
US5541197A (en) * 1994-04-26 1996-07-30 Merck & Co., Inc. Substituted sulfonamides as selective β3 agonists for the treatment of diabetes and obesity
US5561142A (en) * 1994-04-26 1996-10-01 Merck & Co., Inc. Substituted sulfonamides as selective β3 agonists for the treatment of diabetes and obesity
EP0764640A1 (en) * 1995-09-21 1997-03-26 Eli Lilly And Company Selective beta3 adrenergic agonists
US5840738A (en) * 1996-09-05 1998-11-24 Eli Lilly And Company Selective β-3 adrenergic agonists
US6046227A (en) * 1997-12-05 2000-04-04 Eli Lilly And Company Selective β3 adrenergic agonists
US6140352A (en) * 1996-09-05 2000-10-31 Eli Lilly And Company Carbazolyl-substituted ethanolamines as selective β-3 agonists
EP1625134A2 (en) * 2003-05-17 2006-02-15 QUEEN MARY &amp; WESTFIELD COLLEGE Substituted phosphonate fluorescent sensors and use thereof
WO2019185314A1 (en) 2018-03-28 2019-10-03 Unilever Plc Non-aluminium antiperspirant compositions

Families Citing this family (3)

* Cited by examiner, † Cited by third party
Publication number Priority date Publication date Assignee Title
TWI501850B (en) * 2013-04-24 2015-10-01 Rexon Ind Corp Ltd Blade guard assembly for tile cutter
CN103372020A (en) * 2013-07-04 2013-10-30 丁圣雨 Application of Chukrasone A in preparation of medicines for promoting insulin secretion
CN117585653B (en) * 2023-11-13 2024-08-09 湖北三峡实验室 Method for preparing low-cost battery-grade ferric phosphate by multiple precipitation method

Citations (3)

* Cited by examiner, † Cited by third party
Publication number Priority date Publication date Assignee Title
EP0102213A1 (en) * 1982-08-21 1984-03-07 Beecham Group Plc Ethanolamine derivatives, pharmaceutical compositions containing them, and processes for preparing them
EP0262785A1 (en) * 1986-08-29 1988-04-06 Beecham Group Plc N-[2-(4-carboxymethoxyphenyl)-1-methylethyl]-2-hydroxy-2-(3-chlorophenyl)ethane amine, the methyl ester, salt and stereoisomeric forms thereof
EP0455006A2 (en) * 1990-05-04 1991-11-06 American Cyanamid Company Substituted 5-(2-((2-aryl-2-hydroxyethyl)amino)propyl)-1,3-benzodioxoles

Patent Citations (3)

* Cited by examiner, † Cited by third party
Publication number Priority date Publication date Assignee Title
EP0102213A1 (en) * 1982-08-21 1984-03-07 Beecham Group Plc Ethanolamine derivatives, pharmaceutical compositions containing them, and processes for preparing them
EP0262785A1 (en) * 1986-08-29 1988-04-06 Beecham Group Plc N-[2-(4-carboxymethoxyphenyl)-1-methylethyl]-2-hydroxy-2-(3-chlorophenyl)ethane amine, the methyl ester, salt and stereoisomeric forms thereof
EP0455006A2 (en) * 1990-05-04 1991-11-06 American Cyanamid Company Substituted 5-(2-((2-aryl-2-hydroxyethyl)amino)propyl)-1,3-benzodioxoles

Cited By (22)

* Cited by examiner, † Cited by third party
Publication number Priority date Publication date Assignee Title
WO1995004047A1 (en) * 1993-07-31 1995-02-09 Smithkline Beecham Plc 2-benzoheterocyclyloxy or thiopropanolamine derivatives with adreno receptor agonist activity
WO1995007284A1 (en) * 1993-09-09 1995-03-16 Smithkline Beecham Plc Phosphinic acid derivatives with anti-hyper glycemic and/or anti-obesity activity
US5541197A (en) * 1994-04-26 1996-07-30 Merck & Co., Inc. Substituted sulfonamides as selective β3 agonists for the treatment of diabetes and obesity
US5561142A (en) * 1994-04-26 1996-10-01 Merck & Co., Inc. Substituted sulfonamides as selective β3 agonists for the treatment of diabetes and obesity
WO1996004233A1 (en) * 1994-07-29 1996-02-15 Smithkline Beecham Plc Aryloxy and arylthiopropanolamine derivatives useful as beta 3-adrenoreceptor agonists and antagonists of the beta 1 and beta 2-adrenoreceptors and pharmaceutical composition thereof
WO1996004234A1 (en) * 1994-07-29 1996-02-15 Smithkline Beecham Plc Aryloxy andarycthio propanolamine derivatives useful as b3-adrenoreceptor agonists and antagons of the b1 and b2 adrenoreceptors and pharmaceutical composition thereof
US5491134A (en) * 1994-09-16 1996-02-13 Bristol-Myers Squibb Company Sulfonic, phosphonic or phosphiniic acid β3 agonist derivatives
US5939443A (en) * 1995-09-21 1999-08-17 Eli Lilly And Company Selective β3 adrenergic agonists
EP0764640A1 (en) * 1995-09-21 1997-03-26 Eli Lilly And Company Selective beta3 adrenergic agonists
US5786356A (en) * 1995-09-21 1998-07-28 Eli Lilly And Company Selective β3 adrenergic agonists
US6060492A (en) * 1995-09-21 2000-05-09 Eli Lilly And Company Selective β3 adrenergic agonists
US6265581B1 (en) 1995-09-21 2001-07-24 Eli Lilly And Company Selective β3 adrenergic agonists
US6686372B2 (en) 1996-09-05 2004-02-03 Eli Lilly And Company Selective β3 adrenergic agonists
US5840738A (en) * 1996-09-05 1998-11-24 Eli Lilly And Company Selective β-3 adrenergic agonists
US6075040A (en) * 1996-09-05 2000-06-13 Eli Lilly And Company Selective β3 adrenergic agonists
US6140352A (en) * 1996-09-05 2000-10-31 Eli Lilly And Company Carbazolyl-substituted ethanolamines as selective β-3 agonists
US6413991B1 (en) 1996-09-05 2002-07-02 Eli Lilly And Company Selective β3 adrenergic agonists
US7041684B2 (en) 1996-09-05 2006-05-09 Eli Lilly And Company Selective β3 adrenergic agonists
US6046227A (en) * 1997-12-05 2000-04-04 Eli Lilly And Company Selective β3 adrenergic agonists
US6617347B1 (en) 1997-12-05 2003-09-09 Eli Lilly And Company Selective β3 adrenergic agonists
EP1625134A2 (en) * 2003-05-17 2006-02-15 QUEEN MARY &amp; WESTFIELD COLLEGE Substituted phosphonate fluorescent sensors and use thereof
WO2019185314A1 (en) 2018-03-28 2019-10-03 Unilever Plc Non-aluminium antiperspirant compositions

Also Published As

Publication number Publication date
DE69311118T2 (en) 1997-09-25
DE69311118D1 (en) 1997-07-03
CN1101349A (en) 1995-04-12
TW249804B (en) 1995-06-21
GB9215844D0 (en) 1992-09-09
JPH07509458A (en) 1995-10-19
EP0652884A1 (en) 1995-05-17
EP0652884B1 (en) 1997-05-28
MX9304467A (en) 1994-04-29
AU4715193A (en) 1994-02-14
ATE153666T1 (en) 1997-06-15
US5519006A (en) 1996-05-21
ZA935340B (en) 1994-05-16
CA2140916A1 (en) 1994-01-26

Similar Documents

Publication Publication Date Title
EP0652884B1 (en) Phosphonated arylethanolamine compounds with anti-hyperglycemic and/or anti-obesity activity
EP0750617A1 (en) Novel heterocyclic ethanolamine derivatives with beta-adrenoreceptor agonistic activity
FR2596393A1 (en) Derivatives of 3-hydroxy-4-(dihydroxooxophosphorio)butanoic acid, process for their preparation, their application as medicament and the compositions which contain them
GB2198134A (en) Substituted alpha -amino acids, their preparation and pharmaceutical compositions containing them
WO1998022480A1 (en) Phosphorus containing aryloxy or arylthio propanolamine derivatives
JPH10503507A (en) Aryloxy and arylthiopropanolamine derivatives useful as beta3-adrenergic receptor agonists and beta1- and beta2-adrenergic receptor antagonists, and pharmaceutical compositions thereof
EP0170121B1 (en) 2-hydroxy-3-aryloxypropyl or 2-hydroxy-2-benzofuranyl ethyl tertiary amine having an anti-hyperglycaemic and/or anti-obesity activity
EP0711284A1 (en) 2-benzoheterocyclyloxy or thiopropanolamine derivatives with adreno receptor agonist activity
EP0582515A1 (en) Monohydrate of the disodium salt of 4-chlorophenylthiomethylene-bisphosphonic acid, its preparation and pharmaceutical compositions containing it
EP0717746B1 (en) Phosphinic acid derivatives with anti-hyper glycemic and/or anti-obesity activity
US4803293A (en) Tertiary amines
WO1993010074A1 (en) Arylethanolamine derivatives and their use as beta-3-adrenoceptor agonists
EP0139921B1 (en) 2-phenylethylamine derivatives
US2725399A (en) Tertiary dialkylamino propanols
US5134134A (en) Unsaturated aminodicarboxylic acid derivatives
US20080032950A1 (en) Neurologically-Active Compounds
US5475129A (en) Phosphonoalkyl phenylalanine compounds suitably protected for use in peptide synthesis
US3839423A (en) Demethylation process
OA11683A (en) Use of thiahzolidinediones for the treatment of hyperglycaemia.
EP0171760A2 (en) Substituted phenoxypropylaminoethoxyphenoxyacetamides
RU2004106531A (en) Salts of Substituted 1,2,3,4-Tetrahydroquinoline-2-Carboxylic Acid Derivatives
FI59791C (en) ANALOGIFICATION OF FARMING AGAINST PHARMACOLOGICAL ANALYSIS E2 ANALOG OF AV PROSTAGLAND
HU181019B (en) Process for producing s-bracket-plus-bracket closed-alpha-square bracket-bracket-methyl-amino-bracket closed-methyl-square bracket closed-9,10-ethano-antracene-9-bracket-10h-bracket closed-ethanol and acid additional salts
TH55868A (en) A novel form of propyl phosphenic amino acid.

Legal Events

Date Code Title Description
AK Designated states

Kind code of ref document: A1

Designated state(s): AT AU BB BG BR CA CH CZ DE DK ES FI GB HU JP KP KR KZ LK LU MG MN MW NL NO NZ PL PT RO RU SD SE SK UA US VN

AL Designated countries for regional patents

Kind code of ref document: A1

Designated state(s): AT BE CH DE DK ES FR GB GR IE IT LU MC NL PT SE BF BJ CF CG CI CM GA GN ML MR NE SN TD TG

DFPE Request for preliminary examination filed prior to expiration of 19th month from priority date (pct application filed before 20040101)
121 Ep: the epo has been informed by wipo that ep was designated in this application
WWE Wipo information: entry into national phase

Ref document number: 1993917898

Country of ref document: EP

WWE Wipo information: entry into national phase

Ref document number: 2140916

Country of ref document: CA

WWE Wipo information: entry into national phase

Ref document number: 254860

Country of ref document: NZ

WWE Wipo information: entry into national phase

Ref document number: 08374745

Country of ref document: US

WWP Wipo information: published in national office

Ref document number: 1993917898

Country of ref document: EP

REG Reference to national code

Ref country code: DE

Ref legal event code: 8642

WWG Wipo information: grant in national office

Ref document number: 1993917898

Country of ref document: EP