WO1995004530A1 - Endothelin receptor antagonists - Google Patents

Endothelin receptor antagonists Download PDF

Info

Publication number
WO1995004530A1
WO1995004530A1 PCT/US1994/008917 US9408917W WO9504530A1 WO 1995004530 A1 WO1995004530 A1 WO 1995004530A1 US 9408917 W US9408917 W US 9408917W WO 9504530 A1 WO9504530 A1 WO 9504530A1
Authority
WO
WIPO (PCT)
Prior art keywords
hydrogen
alkyl
alkoxy
halogen
compound
Prior art date
Legal status (The legal status is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the status listed.)
Ceased
Application number
PCT/US1994/008917
Other languages
French (fr)
Inventor
John Duncan Elliott
Jia-Ning Xiang
Current Assignee (The listed assignees may be inaccurate. Google has not performed a legal analysis and makes no representation or warranty as to the accuracy of the list.)
SmithKline Beecham Corp
Original Assignee
SmithKline Beecham Corp
Priority date (The priority date is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the date listed.)
Filing date
Publication date
Application filed by SmithKline Beecham Corp filed Critical SmithKline Beecham Corp
Priority to EP94925208A priority Critical patent/EP0711157A4/en
Priority to JP7506557A priority patent/JPH09501178A/en
Publication of WO1995004530A1 publication Critical patent/WO1995004530A1/en
Anticipated expiration legal-status Critical
Ceased legal-status Critical Current

Links

Classifications

    • CCHEMISTRY; METALLURGY
    • C07ORGANIC CHEMISTRY
    • C07DHETEROCYCLIC COMPOUNDS
    • C07D317/00Heterocyclic compounds containing five-membered rings having two oxygen atoms as the only ring hetero atoms
    • C07D317/08Heterocyclic compounds containing five-membered rings having two oxygen atoms as the only ring hetero atoms having the hetero atoms in positions 1 and 3
    • C07D317/44Heterocyclic compounds containing five-membered rings having two oxygen atoms as the only ring hetero atoms having the hetero atoms in positions 1 and 3 ortho- or peri-condensed with carbocyclic rings or ring systems
    • C07D317/46Heterocyclic compounds containing five-membered rings having two oxygen atoms as the only ring hetero atoms having the hetero atoms in positions 1 and 3 ortho- or peri-condensed with carbocyclic rings or ring systems condensed with one six-membered ring
    • C07D317/48Methylenedioxybenzenes or hydrogenated methylenedioxybenzenes, unsubstituted on the hetero ring
    • C07D317/50Methylenedioxybenzenes or hydrogenated methylenedioxybenzenes, unsubstituted on the hetero ring with only hydrogen atoms, hydrocarbon or substituted hydrocarbon radicals, directly attached to atoms of the carbocyclic ring
    • C07D317/60Radicals substituted by carbon atoms having three bonds to hetero atoms with at the most one bond to halogen, e.g. ester or nitrile radicals
    • AHUMAN NECESSITIES
    • A61MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61PSPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
    • A61P13/00Drugs for disorders of the urinary system
    • A61P13/02Drugs for disorders of the urinary system of urine or of the urinary tract, e.g. urine acidifiers
    • AHUMAN NECESSITIES
    • A61MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61PSPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
    • A61P15/00Drugs for genital or sexual disorders; Contraceptives
    • AHUMAN NECESSITIES
    • A61MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61PSPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
    • A61P43/00Drugs for specific purposes, not provided for in groups A61P1/00-A61P41/00
    • AHUMAN NECESSITIES
    • A61MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61PSPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
    • A61P9/00Drugs for disorders of the cardiovascular system
    • AHUMAN NECESSITIES
    • A61MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61PSPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
    • A61P9/00Drugs for disorders of the cardiovascular system
    • A61P9/08Vasodilators for multiple indications
    • AHUMAN NECESSITIES
    • A61MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61PSPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
    • A61P9/00Drugs for disorders of the cardiovascular system
    • A61P9/10Drugs for disorders of the cardiovascular system for treating ischaemic or atherosclerotic diseases, e.g. antianginal drugs, coronary vasodilators, drugs for myocardial infarction, retinopathy, cerebrovascula insufficiency, renal arteriosclerosis
    • AHUMAN NECESSITIES
    • A61MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61PSPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
    • A61P9/00Drugs for disorders of the cardiovascular system
    • A61P9/12Antihypertensives

Definitions

  • the present invention relates to novel compounds, pharmaceutical compositions containing these compounds and their use as endothelin receptor antagonists.
  • Endothelin is a highly potent vasoconstrictor peptide synthesized and released by the vascular endothelium. Endothelin exists as three isoforms, ET-1, ET-2 and ET-3. [Unless otherwise stated "endothelin” shall mean any or all of the isoforms of endothelin]. Endothelin has profound effects on the cardiovascular system, and in particular, the coronary, renal and cerebral circulation. Elevated or abnormal release of endothelin is associated with smooth muscle contraction which is involved in the pathogenesis of cardiovascular, cerebrovascular, respiratory and renal pathophysiology. Elevated levels of endothelin have been reported in plasma from patients with essential hypertension, acute myocardial infarction, subarachnoid hemorrhage, atherosclerosis, and patients with uraemia undergoing dialysis.
  • endothelin has pronounced effects on blood pressure and cardiac output.
  • An intravenous bolus injection of ET (0.1 to 3 nmol/kg) in rats causes a transient, dose-related depressor response (lasting 0.5 to 2 minutes) followed by a sustained, dose-dependent rise in arterial blood pressure which can remain elevated for 2 to 3 hours following dosing. Doses above 3 nmol/kg in a rat often prove fatal.
  • Endothelin appears to produce a preferential effect in the renal vascular bed. It produces a marked, long-lasting decrease in renal blood flow, accompanied by a significant decrease in GFR, urine volume, urinary sodium and potassium excretion. Endothelin produces a sustained antinatriuretic effect, despite significant elevations in atrial natriuretic peptide. Endothelin also stimulates plasma renin activity.
  • ET may be an important mediator of cerebral vasospasm, a frequent and often fatal consequence of subarachnoid hemorrhage. ET also exhibits direct central nervous system effects such as severe apnea and ischemic lesions which suggests that ET may contribute to the development of cerebral infarcts and neuronal death.
  • endothelin has been found to be a potent constrictor of isolated mammalian airway tissue including human bronchus (Uchida et al., Eur J. of Pharm. 154: 227-228 1988, LaGente, Clin. Exp. Allergy 20: 343-348, 1990; and Springall et al., Lancet.337: 697-701, 1991).
  • Endothelin may play a role in the pathogenesis of interstitial pulmonary f ⁇ brosis and associated pulmonary hypertension, Glard et al., Third International Conference on Endothelin, 1993, p. 34 and ARDS (Adult Respiratory Distress Syndrome), Sanai et al. , Supra, p. 112.
  • Endothelin has been associated with the induction of hemorrhagic and necrotic damage in the gastric mucosa (Whittle et al., Br. J. Pharm.95: 1011-1013, 1988); Raynaud's phenomenon, Cinniniello et al., Lancet 337: 114-115, 1991); Crohn's Disease and ulcerative colitis, Munch et al., Lancet. Vol. 339, p. 381; Migraine (Edmeads, Headache, Feb. 1991 p 127); Sepsis (Weitzberg et al., Circ. Shock 33: 222-227, 1991; Pittet et al., Ann. Surg.
  • Endothelin stimulates both bone resorption and anabolism and may have a role in the coupling of bone remodeling. Tatrai et al. Endocrinology. Vol. 131, p. 603-607.
  • Endothelin has been reported to stimulate the transport of sperm in the uterine cavity, Casey et al., J. Clin. Endo and Metabolism. Vol. 74, No. 1, p. 223-225, therefore endothelin antagonists may be useful as male contraceptives.
  • Endothelin modulates the ovarian/menstrual cycle, Kenegsberg, J. of Clin. Endo. and Met., Vol. 74, No. 1, p. 12, and may also play a role in the regulation of penile vascular tone in man, Lau et al., Asia Pacific J. of Pharm., 1991, 6:287-292 and Tejada et al., J. Amer. Physio. Soc. 1991, H1078-H1085.
  • endothelin receptor antagonists would offer a unique approach toward the phaimacotherapy of hypertension, renal failure, cerebrovascular disease, myocardial ischemia, angina, heart failure, asthma, atherosclerosis, Raynaud's phenomenon, ulcers, sepsis, migraine, glaucoma, endotoxin shock, endotoxin induced multiple organ failure or disseminated intravascular coagulation, cyclosporin-induced renal failure and as an adjunct in angioplasty for prevention of restenosis, diabetes, preclampsia of pregnancy, bone remodeling, kidney transplant, male contraceptives, infertility and priaprism.
  • Atherosclerosis and as an adjunct in angioplasty for prevention of restenosis.
  • This invention further constitutes a method for antagonizing endothelin receptors in an animal, including humans, which comprises
  • R 1 is -X(CH 2 ) n Ar or -X(CH 2 ) n R 8 or
  • R 2 is hydrogen, Ar or (c);
  • P 1 is -X(CH 2 ) n R 8 ;
  • P 2 is -X(CH 2 ) n R 8 , or-XR 9 Y;
  • R 3 and R 5 are independently hydrogen, R 11 , OH,
  • -X(CH 2 ) n R 8 may be unsubstituted or substituted by one or two -(CH 2 ) n Ar groups;
  • R4 is hydrogen, R 11 , OH, C 1-5 alkoxy, -S(O) q R 11 , -N(R 6 )2, -X(R 11 ), Br, F, I, Cl or -NHCOR 6 wherein the C 1-8 alkoxy may be unsubstituted or substituted by OH, methoxy or halogen;
  • R 6 is independently hydrogen or C 1-8 alkyl
  • R 7 is independently hydrogen, C 1-10 alkyl. C 2-10 alkeny or C 2-8 alkynyl, all of which may be unsubstituted or substituted by one or more OH, -N(R 6 ) 2 , -CO 2 R 12 , halogen or XC 1-5 alkyl; or R 7 is (CH 2 ) n Ar; R 8 is hydrogen, R 11 , -CO 2 R 7 , -CO 2 C(R 7 ) 2 O(CO)XR 11 ,
  • R 10 is R 3 or R 4 ;
  • R 11 is Ar, C 3-8 cycloalkyl, C 1-8 alkyl, C 2-8 alkenyl, C 2-8 alkynyl all of which may be unsubstituted or substituted by one or more OH, CH 2 OH, N(R 6 ) 2 or halogen;
  • R 12 is hydrogen, C 1-6 alkyl, C 2-6 alkenyl or C 2-7 alkynyl;
  • X is (CH 2 ) n , O, NR 6 or S(O) q ;
  • Y is CH 3 or X(CH 2 ) n Ar
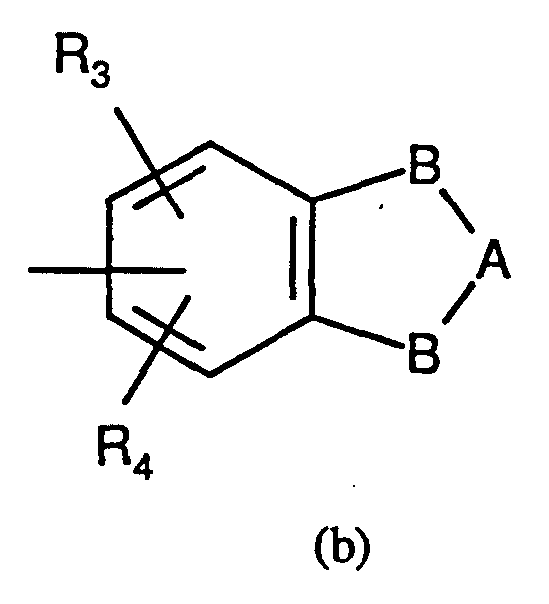
  • Ar is:
  • thiazolidinyl isoxazolyl, oxadiazolyl, thiadiazolyl, morpholinyl, piperidinyl, piperazinyl, pyrrolyl, or pyrimidyl; all of which may be unsubstituted or substituted by one or more R 3 or R 4 groups;
  • B is -CH 2 - or -O-;
  • Z 1 and Z 2 are independently hydrogen, C 1-8 alkyl, C 2-8 alkenyl, C 2- 8 alkynyl, OH, C 1-8 alkoxy, S(O) q C 1-8 alkyl, N(R 6 ) 2 , Br, F, I, Cl, NHCOR 6 , -X(CH 2 ) n R 8 , phenyl, benzyl or C 3-6 cycloalkyl wherein the C 1-8 alkyl, C 2- 8 alkenyl or C 2-8 alkynyl may be optionally substituted by COOH, OH,
  • q is zero, one or two
  • n is an integer from 0 to six;
  • n 1, 2 or 3; and the dotted line indicates the optional presence of one or two double bonds; or a pharmaceutically acceptable salt thereof; provided that:
  • R 2 is not hydrogen when X is S(O) q ;
  • R 1 when an optional double bond is present and R 1 is attached directly to the double bond, R 1 is not NR 6 Ar,
  • R 3 , R 5 , Z 1 , Z 2 , or Z 3 is X(CH 2 ) n R 8 and n is 0, X is not oxygen or NR6 when R 8 is OR 6 or CO 2 H;
  • R 1 and R 2 are phenyl, R 3 , R 4 and R 5 are not all hydrogen.
  • alkyl, alkenyl, alky nyl and alkoxy groups may be straight or branched.
  • halogen is used to mean iodo, fluoro, chloro or bromo.
  • the compounds of the present invention may contain one or more asymmetric carbon atoms and may exist in racemic and optically active form. All of these compounds and diastereoisomers are contemplated to be within the scope of the present invention.
  • R 1 is X(CH 2 ) n Ar, (Ar is (a) or (b)), dihydrobenzofuranyl, benzodioxanyl, cyclohexyl, C 1-4 alkyl
  • R 2 is (a), (b) C 1-4 alkyl, indolyl or hydrogen
  • R 3 and R 5 are independently hydrogen, OH, C 1-5 alkoxy, halogen, -OC 1-4 alkyl phenyl, R CO 2 R 7, C 1-4 alkyl, N(R 6 )2, NH(CO)CH 3, -X(CH 2 ) n R 8 , -XR 9 pyridyl, phenyl or S(O) p C 1-5 alkyl
  • R 4 is hydrogen, OH, C 1-5 alkoxy, halogen, C 1-4 alkyl, N(R 6 ) 2 , NH(CO)CH 3 or
  • Z 1 , Z 2 and Z 3 are independently XR 9 Y, benzyl, hydrogen, OH, C 1-5 alkoxy, -N(R 6 ) 2 , S(O) q C 1-8 alkyl, NHCOR 6 , X(CH 2 ) n R 8 or halogen, or Z 1 and Z 2 together may be -O-A-O on contiguous carbons; P 1 and P 2 are
  • Ar is (a), (b), phenyl, or pyridyl;
  • X is (CH 2 ) n or oxygen.
  • R 3 is hydrogen or - X(CH 2 ) n R 8 , R 11 CO 2 R 7 ; R 4 and R 5 are independently hydrogen, OH,
  • Z 1 and Z 3 are hydrogen and Z 2 is hydrogen, OH, C 1-5 alkoxy, halogen, X(CH 2 ) n R 8 , NH 2 , benzyl, NH(CO)CH 3 , or Z 1 and Z 2 together may be O-A-O.
  • the present invention provides Compounds of Formula (I)
  • Benzylsilanes of Formula (4) may be prepared from the corresponding ortho-methyl benzoic acid esters (6) by treatment with a base such as lithium di- isopropylamide in the presence of chlorotrimethylsilane in a solvent such as anhydrous ether.
  • Saponification of a compound of Formula (5) using aqueous sodium or potassium hydroxide in a suitable solvent such as methanol or isopropanol followed by acidification with a dilute mineral acid such as hydrochloric affords a tri-acid of structure (7).
  • Saponification of an ester of Formula (15) using aqueous sodium or potassium hydroxide in a suitable solvent such as methanol or isopropanol affords, after acidification with a dilute mineral acid such as hydrochloric, an acid of Formula (16).
  • An enol triflate of Formula (17) may also be prepared from a compound of structure (10) by treatment with a base such as N,N-di-isopropylethylamine in a solvent such as dichloromethane followed by trifluoromethanesulfonic anhydride.
  • a base such as N,N-di-isopropylethylamine
  • a solvent such as dichloromethane followed by trifluoromethanesulfonic anhydride.
  • Organocerium compounds of Formula (20) may be prepared by treatment of the corresponding orgnolidiium compound with anhydrous cerium III chloride at -78°C in a solvent such as tetrahydrofuran.
  • Compounds of Formula (I) and their pharmaceutically acceptable salts may be administered in a standard manner for the treatment of the indicated diseases, for example orally, parenterally, sub-lingually, transdermally, rectally, via inhalation or via buccal administration.
  • a syrup formulation will generally consist of a suspension or solution of the compound or salt in a liquid carrier for example, ethanol, peanut oil, olive oil, glycerine or water with a flavouring or colouring agent.
  • a liquid carrier for example, ethanol, peanut oil, olive oil, glycerine or water with a flavouring or colouring agent.
  • any pharmaceutical carrier routinely used for preparing solid formulations may be used. Examples of such carriers include magnesium stearate, terra alba, talc, gelatin, agar, pectin, acacia, stearic acid, starch, lactose and sucrose.
  • composition is in the form of a capsule
  • any routine encapsulation is suitable, for example using the aforementioned carriers in a hard gelatin capsule shell.
  • composition is in the form of a soft gelatin shell capsule
  • any pharmaceutical carrier routinely used for preparing dispersions or suspensions may be considered, for example aqueous gums, celluloses, silicates or oils and are incorporated in a soft gelatin capsule shell.
  • Typical parenteral compositions consist of a solution or suspension of the compound or salt in a sterile aqueous or non-aqueous carrier optionally containing a parenterally acceptable oil, for example polyethylene glycol, polyvinylpyrrolidone, lecithin, arachis oil, or sesame oil.
  • a parenterally acceptable oil for example polyethylene glycol, polyvinylpyrrolidone, lecithin, arachis oil, or sesame oil.
  • compositions for inhalation are in the form of a solution, suspension or emulsion that may be administered as a dry powder or in the form of an aerosol using a conventional propellant such as dichlorodifluoromethane or trichlorofluoromethane.
  • a typical suppository formulation comprises a compound of Formula (I) or a pharmaceutically acceptable salt thereof which is active when administered in this way, with a binding and/or lubricating agent, for example polymeric glycols, gelatins, cocoa-butter or other low melting vegetable waxes or fats or their synthetic analogues.
  • Typical transdermal formulations comprise a conventional aqueous or non-aqueous vehicle, for example a cream, ointment, lotion or paste or are in the form of a medicated plaster, patch or membrane.
  • the composition is in unit dosage form, for example a tablet, capsule or metered aerosol dose, so that the patient may administer to themselves a single dose.
  • Each dosage unit for oral administration contains suitably from 0.1 mg to 500 mg/Kg, and preferably from 1 mg to 100 mg/Kg, and each dosage unit for parenteral administration contains suitably from 0.1 mg to 100 mg, of a compound of Formula (I) or a pharmaceutically acceptable salt thereof calculated as the free acid.
  • Each dosage unit for intranasal administration contains suitably 1-400 mg and preferably 10 to 200 mg per person.
  • a topical formulation contains suitably 0.01 to 1.0% of a compound of Formula (I).
  • the daily dosage regimen for oral administration is suitably about 0.01 mg/Kg to 40 mg/Kg, of a compound of Formula (I) or a pharmaceutically acceptable salt thereof calculated as the free acid.
  • the daily dosage regimen for parenteral administration is suitably about 0.001 mg/Kg to 40 mg/Kg, of a compound of the Formula (I) or a pharmaceutically acceptable salt thereof calculated as the free acid.
  • the daily dosage regimen for intranasal administration and oral inhalation is suitably about 10 to about 500 mg/person.
  • the active ingredient may be administered from 1 to 6 times a day, sufficient to exhibit the desired activity.
  • Rat cerebellum or kidney cortex were rapidly dissected and frozen immediately in liquid nitrogen or used fresh.
  • the tissues 1-2 g for cerebellum or 3-5 g for kidney cortex, were homogenized in 15 mls of buffer containing 20mM Tris HCI and 5mM EDTA, pH 7.5 at 4°C using a motor-driven homogenizer.
  • the homogenates were filtered through cheesecloth and centrifuged at 20,000 x g for 10 minutes at 4°C. The supernatant was removed and centrifuged at 40,000 xg for 30 minutes at 4°C.
  • the resulting pellet was resuspended in a small volume of buffer containing 50 mM Tris, 10 mM MgCl 2 , pH 7.5; aliquotted with small vials and frozen in liquid nitrogen.
  • the membranes were diluted to give 1 and 5 mg of protein for each tube for cerebellum and kidney cortex in the binding assay.
  • Freshly isolated rat mesenteric artery and collateral vascular bed were washed in ice cold saline (on ice) and lymph nodes were removed from along the major vessel. Then, the tissue was homogenized using a polytron in buffer containing 20 mM Tris and 5mM EDTA, pH 7.5 at 4°C in 15 ml volume for ⁇ 6 gm of mesenteric artery bed. The homogenate was strained through cheesecloth and centrifuged at 2,000 xg for 10 min. at 4°C. The supernatant was removed and centrifuged at 40,000 xg for 30 min. at 4°C. The resulting pellet was resuspended as explained above for cerebellum and kidney cortex. Approximately 10 mg of membrane protein was used for each tube in binding experiments.
  • [ 125 I]ET-1 binding to membranes from rat cerebellum (2-5 mg protein/assay tube) or kidney cortex (3-8 mg protein/assay tube) were measured after 60 minutes incubation at 30°C in 50 mM Tris HCI, 10 mM MgCl 2 , 0.05% BSA, pH 7.5 buffer in a total volume of 100 ml.
  • Membrane protein was added to tubes containing either buffer or indicated concentration of compounds.
  • [ 125 I]ET- 1 (2200 Ci/mmol) was diluted in the same buffer containing BSA to give a final concentration of 0.2-0.5 nM ET-1. Total and nonspecific binding were measured in the absence and presence of 100 nM unlabelled ET-1.
  • Rat aorta are cleaned of connective tissue and adherent fat, and cut into ring segments approximately 3 to 4 mm in length.
  • Vascular rings are suspended in organ bath chambers (10 ml) containing Krebs-bicarbonate solution of the following composition (millimolar): NaCI, 112.0; KC1, 4.7; KH 2 PO 4 , 1.2; MgSO 4 , 1.2; CaCl 2 , 2.5; NaHCO 3 , 25.0; and dextrose, 11.0.
  • Tissue bath solutions are maintained at 37°C and aerated continuously with 95% O 2 / 5%° CO 2 .
  • ET-1 induced vascular contractions are expressed as a percentage of the response elicited by 60 mM KCl for each individual tissue which is determined at the beginning of each experiment. Data are expressed as the mean ⁇ S.E.M. Dissociation constants (K b ) of competitive antagonists were determined by the standard method of Arunlakshana and Schild.
  • MeOH/THF was added 2 mL of 10% NaOH and heated to 70 °C for 7 h. The mixture was poured into water and washed with 1:1 Hexane/EtOAc. The aqeous solution was acidified with 6N HCl to pH 5 and extracted with EtOAc. The organic extract was washed eith brine and dried (Na 2 SO 4 ).
  • Formulations for pharmaceutical use incorporating compounds of the present invention can be prepared in various forms and with numerous excipients. Examples of such formulations are given below.
  • a compound of Formula (I) (1 g) is aerosolized from a metered dose inhaler to deliver the desired amount of drug per use.
  • Step 1 Blend ingredients No. 1, No.2, No. 3 and
  • Step 2 Add sufficient water portion-wise to the blend
  • Step 3 The wet mass is converted to granules by
  • Step 4 The wet granules are then dried in an oven at
  • Step 5 The dry granules are lubricated with
  • Step 6 The lubricated granules are compressed on a
  • a pharmaceutical composition for parenteral administration is prepared by dissolving an appropriate amount of a compound of Formula (I) in polyethylene glycol with heating. This solution is then diluted with water for injections Ph Eur. (to 100 ml). The solution is then steriled by filtration through a 0.22 micron membrane filter and sealed in sterile containers.

Landscapes

  • Health & Medical Sciences (AREA)
  • Chemical & Material Sciences (AREA)
  • Organic Chemistry (AREA)
  • Engineering & Computer Science (AREA)
  • Bioinformatics & Cheminformatics (AREA)
  • Life Sciences & Earth Sciences (AREA)
  • Public Health (AREA)
  • General Chemical & Material Sciences (AREA)
  • Medicinal Chemistry (AREA)
  • Nuclear Medicine, Radiotherapy & Molecular Imaging (AREA)
  • Chemical Kinetics & Catalysis (AREA)
  • Pharmacology & Pharmacy (AREA)
  • Veterinary Medicine (AREA)
  • Animal Behavior & Ethology (AREA)
  • General Health & Medical Sciences (AREA)
  • Heart & Thoracic Surgery (AREA)
  • Cardiology (AREA)
  • Urology & Nephrology (AREA)
  • Vascular Medicine (AREA)
  • Endocrinology (AREA)
  • Reproductive Health (AREA)
  • Pharmaceuticals Containing Other Organic And Inorganic Compounds (AREA)
  • Heterocyclic Compounds That Contain Two Or More Ring Oxygen Atoms (AREA)
  • Acyclic And Carbocyclic Compounds In Medicinal Compositions (AREA)

Abstract

Naphthoic derivatives and pharmaceutical compositions containing these compounds, and their use as endothelin receptor antagonists.

Description

ENDOTHELIN RECEPTOR ANTAGONISTS
FIELD OF INVENTION
The present invention relates to novel compounds, pharmaceutical compositions containing these compounds and their use as endothelin receptor antagonists.
Endothelin (ET) is a highly potent vasoconstrictor peptide synthesized and released by the vascular endothelium. Endothelin exists as three isoforms, ET-1, ET-2 and ET-3. [Unless otherwise stated "endothelin" shall mean any or all of the isoforms of endothelin]. Endothelin has profound effects on the cardiovascular system, and in particular, the coronary, renal and cerebral circulation. Elevated or abnormal release of endothelin is associated with smooth muscle contraction which is involved in the pathogenesis of cardiovascular, cerebrovascular, respiratory and renal pathophysiology. Elevated levels of endothelin have been reported in plasma from patients with essential hypertension, acute myocardial infarction, subarachnoid hemorrhage, atherosclerosis, and patients with uraemia undergoing dialysis.
In vivo, endothelin has pronounced effects on blood pressure and cardiac output. An intravenous bolus injection of ET (0.1 to 3 nmol/kg) in rats causes a transient, dose-related depressor response (lasting 0.5 to 2 minutes) followed by a sustained, dose-dependent rise in arterial blood pressure which can remain elevated for 2 to 3 hours following dosing. Doses above 3 nmol/kg in a rat often prove fatal.
Endothelin appears to produce a preferential effect in the renal vascular bed. It produces a marked, long-lasting decrease in renal blood flow, accompanied by a significant decrease in GFR, urine volume, urinary sodium and potassium excretion. Endothelin produces a sustained antinatriuretic effect, despite significant elevations in atrial natriuretic peptide. Endothelin also stimulates plasma renin activity. These findings suggest that ET is involved in the regulation of renal function and is involved in a variety of renal disorders including acute renal failure, cyclosporine nephrotoxicity and chronic renal failure.
Studies have shown that in vivo, the cerebral vasculature is highly sensitive to both the vasodilator and vasoconstrictor effects of endothelin.
Therefore, ET may be an important mediator of cerebral vasospasm, a frequent and often fatal consequence of subarachnoid hemorrhage. ET also exhibits direct central nervous system effects such as severe apnea and ischemic lesions which suggests that ET may contribute to the development of cerebral infarcts and neuronal death.
ET has also been implicated in myocardial ischemia (Nichols et al. Br. J. Pharm. 99: 597-601, 1989 and Clozel and Clozel, Circ. Res., 65: 1193-1200, 1989) coronary vasospasm (Fukuda et al., Eur. J. Pharm. 165: 301-304, 1989 and Lϋscher, Circ. 83: 701, 1991) heart failure, proliferation of vascular smooth muscle cells, (Takagi. Biochem & Biophys. Res. Commun.; 168: 537-543, 1990, Bobek et al., Am. J. Physiol. 258:408-C415, 1990) and atherosclerosis, (Nakaki et al.,
Biochem. & Biophys. Res. Commun. 158: 880-881, 1989, and Lerman et al., New Eng. J. of Med. 325: 997-1001, 1991). Increased levels of endothelin have been shown after coronary balloon angioplasty (Kadel et al., No. 2491 Circ. 82: 627, 1990).
Further, endothelin has been found to be a potent constrictor of isolated mammalian airway tissue including human bronchus (Uchida et al., Eur J. of Pharm. 154: 227-228 1988, LaGente, Clin. Exp. Allergy 20: 343-348, 1990; and Springall et al., Lancet.337: 697-701, 1991). Endothelin may play a role in the pathogenesis of interstitial pulmonary fϊbrosis and associated pulmonary hypertension, Glard et al., Third International Conference on Endothelin, 1993, p. 34 and ARDS (Adult Respiratory Distress Syndrome), Sanai et al. , Supra, p. 112.
Endothelin has been associated with the induction of hemorrhagic and necrotic damage in the gastric mucosa (Whittle et al., Br. J. Pharm.95: 1011-1013, 1988); Raynaud's phenomenon, Cinniniello et al., Lancet 337: 114-115, 1991); Crohn's Disease and ulcerative colitis, Munch et al., Lancet. Vol. 339, p. 381; Migraine (Edmeads, Headache, Feb. 1991 p 127); Sepsis (Weitzberg et al., Circ. Shock 33: 222-227, 1991; Pittet et al., Ann. Surg. 213: 262-264, 1991), Cyclosporin-induced renal failure or hypertension (Eur. J. Pharmacol.. 180: 191-192, 1990. Kidney Int, 37: 1487-1491, 1990) and endotoxin shock and other endotoxin induced diseases (Biochem. Biophys. Res. Commun., 161: 1220-1227, 1989, Acta Physiol. Scand. 137: 317-318, 1989) and inflammatory skin diseases. (Clin Res. 41:451 and 484, 1993).
Endothelin has also been implicated in preclampsia of pregnancy. Clark et al., Am. J. Obstet. Gynecol. March 1992, p. 962-968; Kamor et al., N. Eng. J. of Med., Nov 22, 1990, p. 1486-1487; Dekker et al., Eur J. Ob. and Gyn. and Rep. Bio. 40 (1991) 215-220; Schiff et al., Am. J. Ostet. Gynecol. Feb 1992, p. 624-628; diabetes mellitus, Takahashi et al., Diabetologia 09901 33:306-310; and acute vascular rejection following kidney transplant, Watschinger et al.,
Transplantation Vol. 52, No. 4, pp. 743-746.
Endothelin stimulates both bone resorption and anabolism and may have a role in the coupling of bone remodeling. Tatrai et al. Endocrinology. Vol. 131, p. 603-607.
Endothelin has been reported to stimulate the transport of sperm in the uterine cavity, Casey et al., J. Clin. Endo and Metabolism. Vol. 74, No. 1, p. 223-225, therefore endothelin antagonists may be useful as male contraceptives. Endothelin modulates the ovarian/menstrual cycle, Kenegsberg, J. of Clin. Endo. and Met., Vol. 74, No. 1, p. 12, and may also play a role in the regulation of penile vascular tone in man, Lau et al., Asia Pacific J. of Pharm., 1991, 6:287-292 and Tejada et al., J. Amer. Physio. Soc. 1991, H1078-H1085.
Thus, endothelin receptor antagonists would offer a unique approach toward the phaimacotherapy of hypertension, renal failure, cerebrovascular disease, myocardial ischemia, angina, heart failure, asthma, atherosclerosis, Raynaud's phenomenon, ulcers, sepsis, migraine, glaucoma, endotoxin shock, endotoxin induced multiple organ failure or disseminated intravascular coagulation, cyclosporin-induced renal failure and as an adjunct in angioplasty for prevention of restenosis, diabetes, preclampsia of pregnancy, bone remodeling, kidney transplant, male contraceptives, infertility and priaprism.
SUMMARY OF THE INVENTION This invention comprises novel compounds represented by Formula
(I) and pharmaceutical compositions containing these compounds, and their use as endothelin receptor antagonists which are useful in the treatment of a variety of cardiovascular and renal diseases including but not limited to: hypertension, acute and chronic renal failure, cyclosporine induced nephrotoxicity, stroke,
cerebrovascular vasospasm, myocardial ischemia, angina, heart failure,
atherosclerosis, and as an adjunct in angioplasty for prevention of restenosis.
This invention further constitutes a method for antagonizing endothelin receptors in an animal, including humans, which comprises
administering to an animal in need thereof an effective amount of a compound of Formula (I). DETAILED DESCRIPTION OF THE INVENTION
The compounds of this invention are represented by structural
Formula (I):
1) A compound of Formula (I),
Figure imgf000006_0001
wherein:
R1 is -X(CH2)nAr or -X(CH2)nR8 or
;
Figure imgf000006_0002
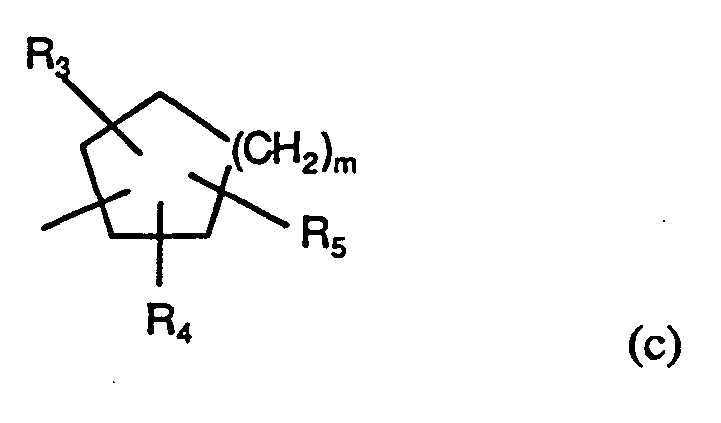
R2 is hydrogen, Ar or (c);
P1 is -X(CH2)nR8;
P2 is -X(CH2)nR8, or-XR9Y;
R3 and R5 are independently hydrogen, R11, OH,
C1-8alkoxy, -S(O)qR11, -N(R6)2, Br, F, I, Cl, CF3, -NHCOR6, -R11CO2R7, -R11CO2R7, -XR9-Y or -X(CH2)nR8 wherein each methylene group within
-X(CH2)nR8 may be unsubstituted or substituted by one or two -(CH2)nAr groups;
R4 is hydrogen, R11, OH, C1-5alkoxy, -S(O)qR11, -N(R6)2, -X(R11), Br, F, I, Cl or -NHCOR6 wherein the C1-8alkoxy may be unsubstituted or substituted by OH, methoxy or halogen;
R6 is independently hydrogen or C1-8alkyl;
R7 is independently hydrogen, C1-10alkyl. C2-10alkeny or C2-8 alkynyl, all of which may be unsubstituted or substituted by one or more OH, -N(R6)2, -CO2R12, halogen or XC1-5 alkyl; or R7 is (CH2)nAr; R8 is hydrogen, R11, -CO2R7, -CO2C(R7)2O(CO)XR11,
PO3(R7)2, -SO2NR7R11, -CONR7SO2R11, -SO3R7, -SO2R7, -P(O)(OR7)R7, CN, -C(O)N(R6)2, -NR7SO2R11, tetrazole or OR6;
R9 is (CH2)n, C1-10alkyl, C2-10alkenyl or phenyl all of which may be unsubstituted or substituted by one or more OH, N(R6)2, COOH, halogen, >C=O or XC1-5alkyl;
R10 is R3 or R4;
R11 is Ar, C3-8cycloalkyl, C1-8alkyl, C2-8alkenyl, C2-8alkynyl all of which may be unsubstituted or substituted by one or more OH, CH2OH, N(R6)2 or halogen;
R12 is hydrogen, C1-6alkyl, C2-6alkenyl or C2-7alkynyl;
X is (CH2)n, O, NR6 or S(O)q;
Y is CH3 or X(CH2)nAr,
Ar is:
Figure imgf000007_0001
Figure imgf000007_0002
naphthyl, indolyl, pyridyl, thienyl, oxazolidinyl, oxazolyl, thiazolyl, isothiazolyl, pyrazolyl, triazolyl, tetrazolyl, imidazolyl, imidazolidinyl,
thiazolidinyl, isoxazolyl, oxadiazolyl, thiadiazolyl, morpholinyl, piperidinyl, piperazinyl, pyrrolyl, or pyrimidyl; all of which may be unsubstituted or substituted by one or more R3 or R4 groups;
A is C=O, or [C(R6)2]m;
B is -CH2- or -O-;
Z1 and Z2 are independently hydrogen, C1-8alkyl, C2-8alkenyl, C2- 8alkynyl, OH, C1-8alkoxy, S(O)qC1-8alkyl, N(R6)2, Br, F, I, Cl, NHCOR6, -X(CH2)nR8, phenyl, benzyl or C3-6cycloalkyl wherein the C1-8alkyl, C2- 8alkenyl or C2-8alkynyl may be optionally substituted by COOH, OH,
CO(CH2)nCH3, CO(CH2)nCH2N(R6)2. or halogen; or Z1 and Z2 together may be -O-A-O- on contiguous carbons; Z3 is Z1 or XR9Y;
q is zero, one or two;
n is an integer from 0 to six;
m is 1, 2 or 3; and the dotted line indicates the optional presence of one or two double bonds; or a pharmaceutically acceptable salt thereof; provided that:
R2 is not hydrogen when X is S(O)q;
when an optional double bond is present there is only one R10, and when the double bond is adjacent to P1 and P2, there is no P1 and P2 is not NR6R9Y;
when an optional double bond is present in Formula (I) and X-R2 is attached to the double bond, X is not NR6;
when an optional double bond is present and R1 is attached directly to the double bond, R1 is not NR6Ar,
when R3, R5, Z1, Z2, or Z3 is X(CH2)nR8 and n is 0, X is not oxygen or NR6 when R8 is OR6 or CO2H;
when R8 is CO(CR11)2O(CO)XR7, X is not S(O)q;
when there are two double bonds present and R1 and R2 are phenyl, R3, R4 and R5 are not all hydrogen.
Also included in the invention are pharmaceutically acceptable salt complexes.
All alkyl, alkenyl, alky nyl and alkoxy groups may be straight or branched. The term "halogen" is used to mean iodo, fluoro, chloro or bromo.
The compounds of the present invention may contain one or more asymmetric carbon atoms and may exist in racemic and optically active form. All of these compounds and diastereoisomers are contemplated to be within the scope of the present invention.
Preferred compounds are those wherein R1 is X(CH2)nAr, (Ar is (a) or (b)), dihydrobenzofuranyl, benzodioxanyl, cyclohexyl, C1-4alkyl; R2 is (a), (b) C1-4alkyl, indolyl or hydrogen; R3 and R5 are independently hydrogen, OH, C1-5alkoxy, halogen, -OC1-4alkyl phenyl, R CO2R7, C1-4alkyl, N(R6)2, NH(CO)CH3, -X(CH2)nR8, -XR9 pyridyl, phenyl or S(O)pC1-5alkyl; R4 is hydrogen, OH, C1-5alkoxy, halogen, C1-4alkyl, N(R6)2, NH(CO)CH3 or
S(O)pC1-5alkyl; Z1, Z2 and Z3 are independently XR9Y, benzyl, hydrogen, OH, C1-5alkoxy, -N(R6)2, S(O)qC1-8alkyl, NHCOR6, X(CH2)nR8 or halogen, or Z1 and Z2 together may be -O-A-O on contiguous carbons; P1 and P2 are
independently hydrogen, CO2H or tetrazole; Ar is (a), (b), phenyl, or pyridyl; X is (CH2)n or oxygen.
More preferred are compounds wherein R3 is hydrogen or - X(CH2)nR8, R11CO2R7; R4 and R5 are independently hydrogen, OH,
C1-5alkoxy, SC1-5alkyl, F, Br, C1-3alkyl or NH2; Z1 and Z3 are hydrogen and Z2 is hydrogen, OH, C1-5alkoxy, halogen, X(CH2)nR8, NH2, benzyl, NH(CO)CH3, or Z1 and Z2 together may be O-A-O.
Most preferred are compounds wherein R1 is (a) or (b) and R2 is (a) or (b); A is CH2, B is -O-; there is no optional double bond; R1 and XR2 are trans to P1; Z2 is OH, C1-5alkoxy, -OCH2CHCH2 or hydrogen, Z1 and Z3 are hydrogen; R3 is XAr, hydrogen, X(CH2)qCOOH, X(CH2)qCONR7SO2R11 or CH=CHCO2H, R4 is hydrogen, substituted phenyl, or C1-2alkoxy; and R5, R10 and P2 are hydrogen.
The present invention provides Compounds of Formula (I)
Figure imgf000009_0001
which for compounds wherein X is (CH2)n, n is 0 and R10 is H, can be prepared by a process which comprises:
a) reacting a substituted aldehyde of Formula (2)
R2 CHO
(2) with dimethyl malonate in a suitable solvent such as benzene with a catalyst such as piperidinium acetate at reflux to provide a compound of Formula (3).
Figure imgf000010_0001
Reaction of a compound of Formula (3) with a benzylsilane of structure (4) (wherein Y is C1-5alkyl)
Figure imgf000010_0002
in the presence of anhydrous cesium fluoride in a solvent such as HMPA at 40-100°C affords compounds of structure (5).
Figure imgf000010_0003
Benzylsilanes of Formula (4) may be prepared from the corresponding ortho-methyl benzoic acid esters (6) by treatment with a base such as lithium di- isopropylamide in the presence of chlorotrimethylsilane in a solvent such as anhydrous ether.
Figure imgf000011_0001
Saponification of a compound of Formula (5) using aqueous sodium or potassium hydroxide in a suitable solvent such as methanol or isopropanol followed by acidification with a dilute mineral acid such as hydrochloric affords a tri-acid of structure (7).
Figure imgf000011_0002
Thermolysis of a compound of Formula (7) at between 150-250°C affords a di-acid of Formula (8).
Figure imgf000011_0003
Re-esterification of a compound of structure (8) with diazomethane in a solvent such as ether or alternatively treatment of (8) with a base such as 1,8-diaza-bicyclo[5.4.0]undec-7-ene in a solvent such as acetonitrile followed by the addition of iodomethane affords diesters of Formula (9). As a further alternative, compounds of structure (9) may be prepared from (8) by treatment with methanolic hydrogen chloride.
Figure imgf000012_0001
Dieckmann cyclization of a di-ester of formula (9) using sodium metfioxide in methanol as a solvent affords, after acidic work-up, a β-keto-ester of Formula (10).
Figure imgf000012_0002
Treatment of a compound of Formula (10) with diazomethane in a solvent such as etiier affords enol ethers of Formula (11).
Figure imgf000012_0003
Reaction of a compound of Formula (11) with a Grignard reagent of Formula (12). R1 MgHaI (12) wherein R1 is X(CH2)nAr, and X is (CH2)n in a solvent such as ether or ether/tetrahydrofuran at approximately 0°C affords compounds of Formula (13).
Figure imgf000013_0001
Hydrogenation of a compound of Formula (13) under hydrogen gas at
approximately 50 p.s.i. in a suitable solvent such as ethyl acetate in the presence of a suitable catalyst such as 5-10% palladium on charcoal affords a racemic compound of Formula (14) with the predominant relative stereochemistry shown.
Figure imgf000013_0002
Epimerization using a base such as potassium t-butoxide in a solvent such as tertiary butanol affords a compound of Formula (15).
Figure imgf000013_0003
Saponification of an ester of Formula (15) using aqueous sodium or potassium hydroxide in a suitable solvent such as methanol or isopropanol affords, after acidification with a dilute mineral acid such as hydrochloric, an acid of Formula (16).
Figure imgf000014_0001
b) As an alternative, treatment of a β-keto-ester of Formula (10) with a base such as sodium hydride in a solvent such as tetrahydrofuran followed by the addition of N-phenyltrifluoromethanesulfonimide affords compounds of Formula (17).
Figure imgf000014_0002
An enol triflate of Formula (17) may also be prepared from a compound of structure (10) by treatment with a base such as N,N-di-isopropylethylamine in a solvent such as dichloromethane followed by trifluoromethanesulfonic anhydride.
Treatment of an enol triflate of Formula (17) with an organostannane of Formula (18)
R 1SnY3
(18) wherein R1 is Ar in a solvent such as dioxane in the presence of a catalyst such as tetrakis(triphenylphosphine)palladium (0) affords compounds of Formula (19).
Figure imgf000015_0001
Hydrogenation of a compound of structure (19) as described above for compounds of Formula (13) affords tetrahydronaphthalenes of Formula (14).
c) As a further alternative treatment of a β-keto-ester of Formula (10) with an organocerium reagent of Formula (20)
(R1 )CeCl2
(20) in a solvent such as tetrahydrofuran at -78°C affords a product of Formula (21). Organocerium compounds of Formula (20) may be prepared by treatment of the corresponding orgnolidiium compound with anhydrous cerium III chloride at -78°C in a solvent such as tetrahydrofuran.
Figure imgf000015_0002
Hydrogenation of compounds of Formula (21) under hydrogen gas at
approximately 50 p.s.i. in a suitable solvent such as ethyl acetate containing 5-20% acetic acid in the presence of a suitable catalyst such as 5-10% palladium on charcoal affords, after saponification as described for compounds of Formula (15), tetrahydronaphthalene carboxyclic acids of Formula (16). d) Saponification of esters of type (13) using aqueous sodium or potassium hydroxide in a suitable solvent such as methanol or isopropanol affords, after acidification with a dilute mineral acid such as hydrochloric, acids of Formula (21), the double bond isomers of which may be separated chromatographically.
With appropriate manipulation and protection of any chemical functionalities, synthesis of the remaining compounds of the Formula (I) is accomplished by methods analogous to those above and to those described in the Experimental section.
In order to use a compound of the Formula (I) or a pharmaceutically acceptable salt thereof for the treatment of humans and other mammals it is normally formulated in accordance with standard pharmaceutical practice as a pharmaceutical composition.
Compounds of Formula (I) and their pharmaceutically acceptable salts may be administered in a standard manner for the treatment of the indicated diseases, for example orally, parenterally, sub-lingually, transdermally, rectally, via inhalation or via buccal administration.
Compounds of Formula (I) and their pharmaceutically acceptable salts which are active when given orally can be formulated as syrups, tablets, capsules and lozenges. A syrup formulation will generally consist of a suspension or solution of the compound or salt in a liquid carrier for example, ethanol, peanut oil, olive oil, glycerine or water with a flavouring or colouring agent. Where the composition is in the form of a tablet, any pharmaceutical carrier routinely used for preparing solid formulations may be used. Examples of such carriers include magnesium stearate, terra alba, talc, gelatin, agar, pectin, acacia, stearic acid, starch, lactose and sucrose. Where the composition is in the form of a capsule, any routine encapsulation is suitable, for example using the aforementioned carriers in a hard gelatin capsule shell. Where the composition is in the form of a soft gelatin shell capsule any pharmaceutical carrier routinely used for preparing dispersions or suspensions may be considered, for example aqueous gums, celluloses, silicates or oils and are incorporated in a soft gelatin capsule shell.
Typical parenteral compositions consist of a solution or suspension of the compound or salt in a sterile aqueous or non-aqueous carrier optionally containing a parenterally acceptable oil, for example polyethylene glycol, polyvinylpyrrolidone, lecithin, arachis oil, or sesame oil.
Typical compositions for inhalation are in the form of a solution, suspension or emulsion that may be administered as a dry powder or in the form of an aerosol using a conventional propellant such as dichlorodifluoromethane or trichlorofluoromethane. A typical suppository formulation comprises a compound of Formula (I) or a pharmaceutically acceptable salt thereof which is active when administered in this way, with a binding and/or lubricating agent, for example polymeric glycols, gelatins, cocoa-butter or other low melting vegetable waxes or fats or their synthetic analogues.
Typical transdermal formulations comprise a conventional aqueous or non-aqueous vehicle, for example a cream, ointment, lotion or paste or are in the form of a medicated plaster, patch or membrane.
Preferably the composition is in unit dosage form, for example a tablet, capsule or metered aerosol dose, so that the patient may administer to themselves a single dose.
Each dosage unit for oral administration contains suitably from 0.1 mg to 500 mg/Kg, and preferably from 1 mg to 100 mg/Kg, and each dosage unit for parenteral administration contains suitably from 0.1 mg to 100 mg, of a compound of Formula (I) or a pharmaceutically acceptable salt thereof calculated as the free acid. Each dosage unit for intranasal administration contains suitably 1-400 mg and preferably 10 to 200 mg per person. A topical formulation contains suitably 0.01 to 1.0% of a compound of Formula (I).
The daily dosage regimen for oral administration is suitably about 0.01 mg/Kg to 40 mg/Kg, of a compound of Formula (I) or a pharmaceutically acceptable salt thereof calculated as the free acid. The daily dosage regimen for parenteral administration is suitably about 0.001 mg/Kg to 40 mg/Kg, of a compound of the Formula (I) or a pharmaceutically acceptable salt thereof calculated as the free acid. The daily dosage regimen for intranasal administration and oral inhalation is suitably about 10 to about 500 mg/person. The active ingredient may be administered from 1 to 6 times a day, sufficient to exhibit the desired activity.
No unacceptable toxicological effects are expected when compounds of the invention are administered in accordance with the present invention.
The biological activity of the compounds of Formula (I) are demonstrated by the following tests:
I. Binding Assay
A) Membrane Preparation
Rat cerebellum or kidney cortex were rapidly dissected and frozen immediately in liquid nitrogen or used fresh. The tissues, 1-2 g for cerebellum or 3-5 g for kidney cortex, were homogenized in 15 mls of buffer containing 20mM Tris HCI and 5mM EDTA, pH 7.5 at 4°C using a motor-driven homogenizer. The homogenates were filtered through cheesecloth and centrifuged at 20,000 x g for 10 minutes at 4°C. The supernatant was removed and centrifuged at 40,000 xg for 30 minutes at 4°C. The resulting pellet was resuspended in a small volume of buffer containing 50 mM Tris, 10 mM MgCl2, pH 7.5; aliquotted with small vials and frozen in liquid nitrogen. The membranes were diluted to give 1 and 5 mg of protein for each tube for cerebellum and kidney cortex in the binding assay.
Freshly isolated rat mesenteric artery and collateral vascular bed were washed in ice cold saline (on ice) and lymph nodes were removed from along the major vessel. Then, the tissue was homogenized using a polytron in buffer containing 20 mM Tris and 5mM EDTA, pH 7.5 at 4°C in 15 ml volume for ~6 gm of mesenteric artery bed. The homogenate was strained through cheesecloth and centrifuged at 2,000 xg for 10 min. at 4°C. The supernatant was removed and centrifuged at 40,000 xg for 30 min. at 4°C. The resulting pellet was resuspended as explained above for cerebellum and kidney cortex. Approximately 10 mg of membrane protein was used for each tube in binding experiments.
B) [125I]ET-1 Binding Protocol
[125I]ET-1 binding to membranes from rat cerebellum (2-5 mg protein/assay tube) or kidney cortex (3-8 mg protein/assay tube) were measured after 60 minutes incubation at 30°C in 50 mM Tris HCI, 10 mM MgCl2, 0.05% BSA, pH 7.5 buffer in a total volume of 100 ml. Membrane protein was added to tubes containing either buffer or indicated concentration of compounds. [125I]ET- 1 (2200 Ci/mmol) was diluted in the same buffer containing BSA to give a final concentration of 0.2-0.5 nM ET-1. Total and nonspecific binding were measured in the absence and presence of 100 nM unlabelled ET-1. After the incubation, the reactions were stopped with 3.0 ml cold buffer containing 50 mM Tris and 10 mM MgCl2, pH 7.5. Membrane bound radioactivity was separated from free ligand by filtering through Whatman GF/C filter paper and washing the filters 5 times with 3 ml of cold buffer using a Brandel cell harvester. Filter papers were counted in a gamma counter with an efficiency of 75%.
II. In Vitro Vascular Smooth Muscle Activity
Rat aorta are cleaned of connective tissue and adherent fat, and cut into ring segments approximately 3 to 4 mm in length. Vascular rings are suspended in organ bath chambers (10 ml) containing Krebs-bicarbonate solution of the following composition (millimolar): NaCI, 112.0; KC1, 4.7; KH2PO4, 1.2; MgSO4, 1.2; CaCl2, 2.5; NaHCO3, 25.0; and dextrose, 11.0. Tissue bath solutions are maintained at 37°C and aerated continuously with 95% O2/ 5%° CO2. Resting tensions of aorta are maintained at 1 g and allowed to equilibrate for 2 hrs., during which time the bathing solution is changed every 15 to 20 min. Isometric tensions are recorded on Beckman R-611 dynographs with Grass FT03 force-displacement transducer. Cumulative concentration-response curves to ET-1 or other contractile agonists are constructed by the method of step- wise addition of the agonist. ET-1 concentrations are increased only after the previous concentration produces a steady-state contractile response. Only one concentration-response curve to ET-1 is generated in each tissue. ET receptor antagonists are added to paired tissues 30 min prior to the initiation of the concentration-response to contractile agonists.
ET-1 induced vascular contractions are expressed as a percentage of the response elicited by 60 mM KCl for each individual tissue which is determined at the beginning of each experiment. Data are expressed as the mean ± S.E.M. Dissociation constants (Kb) of competitive antagonists were determined by the standard method of Arunlakshana and Schild.
The following examples are illustrative and are not limiting of the compounds of this invention.
EXAMPLE 1
(1RS, 2SR, 3SR)-1,3-Bis(3,4-methylene-dioxyphenyl)-1,2,3,4-tetrahydro-2-naphthoic acid a) Benzyl-methyl-2-[1-(3,4-methylenedioxyphenyl)-2-(2- methylcarboxyphenyl)ethyl] malonate.
To a solution of benzyl, methyl-2-(3,4-methylenedioxybenzyliden)-malonate (2.16 g, 6.35 mmol) and metiiyl o-trimethylsilylmethyl benzoate (2.12 g, 9.53 mmol) in HMPA (6 mL) at room temperature was added CsF (1.93 g, 12.70 mmol) in one portion. The mixture was heated to 60 °C for 3 h under argon and then quenched with water, and extracted with 1:1 EtOAc/Hexane. The organic layer was separated and washed with brine and dried (NaSO4). After removing the solvent, chromatography of the residue with 4: 1 Hexane/EtOAc gave 1.87 g (60%) of an inseparable 1:1 mixture of title compounds as colorless oil: Rf 0.31 (silica gel, 2:1 n-hexane/EtOAc). b) Methyl-3-(3,4-methylenedioxyphenyl)-4-(2-methylcarboxyphenyl)butanoate. To a solution of triester (a) (3.30 g, 6.73 mmol) in 10 mL of ether acetate was added 330 mg of 10% Pd/C and the mixture was stirred under H2 atmosphere at room temperature for 7 h. Filtration and concentration gave 2.70 g of the crude acids which were directly used in the following reaction without further purification.
The neat crude acids (2.70 g) were heated to 150 °C for 1 h and flash chromatography of the residue with 4:1 n-hexane/ethyl acetate gave 2.30 g (96% over two steps) of (b) as a colorless oil: Rf 0.46 (silica gel, 2: 1 n- hexane/EtOAc). c) (±)-Phenethyl╌(3SR)-1-methoxy-3-(3,4-methylenedioxyphenyl)-3,4-dihydro-2- naphthoate.
To a solution of diester (800 mg, 2.24 mmol) in 10 mL of THF was added 3 mL of a solution of 25% wt NaOMe/MeOH and the mixture was heated to reflux for 1 h. After cooling to room temperature, the mixture was poured into IN HCI and extracted twice with 1:1 hexane/EtOAc. The combined oraginc extract was washed with brine and dried (Na2SO4). Removal of the solvent gave the crude b-keto ester, which was used in the next reaction without further purification.
To a solution of the crude b-keto ester in 10 mL of toluene was added phenetiiyl alcohol (0.41 mL, 3.36 mmol) and DMAP (10 mg) and the mixture was refluxed for 28 h. After cooling to room temperature, the mixture was poured into water and extracted twice with 1:1 hexane/EtOAc. The combined oraginc extract was washed with brine and dried (Na2SO4). Removal of the solvent gave the crude b-keto ester, which was used in the next reaction without further purification. To a solution of the crude ester in 10 mL of ethanol at room temperature was added 20 mL of diazomediane solution (13 mmol) and the mixture was stirred under argon at room temperature for 17 h. The reaction was quenched with AcOH and the solution was then concentrated. Flash chromatography of the residue with 4:1 hexane/EtOAc gave 647 mg (67% over 3 steps) of a 1:1 mixture of inseparable diastereomeric methyl enoethers (c) as colorless oil. d) (±)-Phenethyl~(3SR)-1,3-bis(3,4-methylenedioxyphenyl)-3,4-dihydro-2- naphthoate.
To a mixture of l-bromo-3,4-methylenedoxybenzene (423 mg, 2.10mmol) and Mg (68 mg, 2.80 mmol) in 3 mL of THF was added 2 uL of Mel and the mixture was irradiated by ultrasound for 30 min. (Mg disappeared). To a solution of methyl enol ether 7 (300 mg, 0.70 mmol) in 4 mL of THF at 0°C under argon was dropwise added the grinard reagent
methylenedioxyphenyl MgBr in THF. After 15 minute, the reaction was quenched with aq. HCI and the mixture was extracted with 1:1 hexane/EtOAc. The organic extract was washed with brine and dried (Na2SO4). After removing the solvent, flash chromatography of the residue with 4: 1
hexane/EtOAc gave 250 mg (69%) of 1:1 mixture of inseparable diastereomers (d) as colorless oil: Rf 0.45 (silica gel, 3:1 n-hexane/EtOAc); MS m/e (MH+) 519; Anal. Calcd. for C33H26O6. C, 76.43; H, 5.05. Found: C, 76.70; H, 5.14. e) (1RS, 2SR, 3SR)-Methyl-1,3-bis(3,4-methylenedioxyphenyl)-1,2,3,4- tetrahydro-2-naphthoate.
To a solution of phenethylester (d) (454 mg, 0.876 mmol) in 4 mL of EtOAc was added 60 mg of 10% Pd/C and shaken under H2 (55 Psi) at room temperature for 24 h. After filtration and concentration, the residue was dissolved in ether and added excess of diazomethane solution in ether. The reaction was quenched with acetic acid and washed with water, brine and dried (Na2SO4). After removing the solvent, flash chromatography of the residue with 4:1 Hexane/EtOAc gave 160 mg (43%) of methyl ester (e) as a colorless oil:Rf 0.52 (silica gel, 3:1 n-hexane/EtOAc). f) (1RS, 2RS, 3SR)-Methyl-1,3-bis(3,4-methylenedioxyphenyl)-1,2,3,4- tetrahydro-2-naphtiιoate.
To solution of methyl ester (e) (160 mg, 0.373 mmol) in 3 mL of THF was added 0.7 mL of 25% NaOMe/MeOH and stirred at room temperature under argon for 18 h. The mixture was poured into 1N HCI and extracted with 1:1 Hexane/EtOAc. The organic extract was washed with brine and dried
(Na2SO4). After removing the solvent, flash chromatography of the residue with 4:1 n-hexane/EtOAc gave 130 mg (81%) of (f) as a white solid: Rf0.52 (silica gel, 3:1 n-hexane/EtOAc). g) (1RS, 2SR, 3SR)-1,3-Bis(3,4-methylenedioxyphenyl)-1,2,3,4-tetrahydro-2- naphthoic acid.
To a solution of methyl ester (f) (110 mg, 0.257 mmol) in 6 mL of 1 : 1
MeOH/THF was added 2 mL of 10% NaOH and heated to 70 °C for 7 h. The mixture was poured into water and washed with 1:1 Hexane/EtOAc. The aqeous solution was acidified with 6N HCl to pH 5 and extracted with EtOAc. The organic extract was washed eith brine and dried (Na2SO4). Removal of the solvent gave 100 mg (94%) of the title acid as a white solid: Rf 0.07 (silica gel, 2:1 n-hexane/EtOAc); 1H NMR (CDCl3) d 3.00-3.25 (m, 3H), 4.38 (d, 7=10.8 Hz, 1H), 5.94 (s, 4H), 6.55-7.18 (m, 10H); MS m/e 416 (M+); FAB HRMS
Calcd. for C25H19O6Na2:461.0977. Found: 461.0954. Anal. Calcd. for C25H19O6Η2O: C, 69.12; H, 5.10. Found: C, 68.89 H, .5.41
EXAMPLE 2
Figure imgf000022_0001
(1RS, 2RS, 3SR)-3-(2-carboxymethoxy-4-methoxyphenyl)-1-(3,4-methylenedioxyphenyl)-1,2,3,4-tetrahydro-2-naphthoic acid.
(1RS, 2SR, 3RS )-1-(2-Carboxymethoxy-4-methoxyphenyl)-3-(3,4-methylenedioxyphenyl)-1,2,3,4-tetrahydro-2-naphthoic acid.
EXAMPLE 4
Formulations for pharmaceutical use incorporating compounds of the present invention can be prepared in various forms and with numerous excipients. Examples of such formulations are given below.
Inhalant Formulation
A compound of Formula (I) (1 g) is aerosolized from a metered dose inhaler to deliver the desired amount of drug per use.
Tablets/Ingredients Per Tablet
1. Active ingredient 40 mg
(Cpd of Form. (I)
2. Corn Starch 20 mg
3. Alginic acid 20 mg
4. Sodium Alginate 20 mg
5. Mg stearate 1.3 mg
2.3 mg Procedure for tablets:
Step 1 Blend ingredients No. 1, No.2, No. 3 and
No.4 in a suitable mixer/blender.
Step 2 Add sufficient water portion-wise to the blend
from Step 1 with careful mixing after each
addition. Such additions of water and mixing
until the mass is of a consistency to permit
its conversion to wet granules.
Step 3 The wet mass is converted to granules by
passing it through an oscillating granulator
using a No. 8 mesh (2.38 mm) screen.
Step 4 The wet granules are then dried in an oven at
140°F (60°C) until dry.
Step 5 The dry granules are lubricated with
ingredient No. 5.
Step 6 The lubricated granules are compressed on a
suitable tablet press.
Parenteral Formulation
A pharmaceutical composition for parenteral administration is prepared by dissolving an appropriate amount of a compound of Formula (I) in polyethylene glycol with heating. This solution is then diluted with water for injections Ph Eur. (to 100 ml). The solution is then steriled by filtration through a 0.22 micron membrane filter and sealed in sterile containers.

Claims

CLAIMS:
1) A compound of Formula (I),
Figure imgf000025_0001
wherein:
R1 is -X(CH2)nAr or -X(CH2)nR8 or
;
Figure imgf000025_0002
R2 is hydrogen, Ar or (c);
P1 is -X(CH2)nR8;
P2 is -X(CH2)nR8, or -XR9Y;
R3 and R5 are independentiy hydrogen, R11, OH,
C1-8alkoxy, -S(O)qR11, N(R6)2, Br, F, I, Cl, CF3, -NHCOR6, -R11CO2R7, -R11CO2R7, -XR9-Y or -X(CH2)nR8 wherein each methylene group within -X(CH2)nR8 may be unsubstituted or substituted by one or two -(CH2)nAr groups;
R4 is hydrogen, R11, OH, C1-5alkoxy, -S(O)qR11, -N(R6)2, -X(R11), Br, F, I, Cl or -NHCOR6 wherein the C1-5alkoxy may be unsubstituted or substituted by OH, methoxy or halogen;
R6 is independentiy hydrogen or C1-4alkyl;
R7 is independently hydrogen, C1-10alkyl, C2-10alkenyl or C2-8 alkynyl, all of which may be unsubstituted or substituted by one or more OH, -N(R6)2, -CO2R12, halogen or XC1-5 alkyl; or R7 is (CH2)nAr;
R8 is hydrogen, R11, -CO2R7, -CO2C(R7)2O(CO)XR11,
PO3(R7)2, -SO2NR7R11, -CONR7SO2R11, -SO3R7, -SO2R7, -P(O)(OR7)R7, CN, -C(O)N(R6)2, -NR7SO2R11, tetrazole or OR6; R9 is (CH2)n, C1-10alkyl, C2-10alkenyl or phenyl all of which may be unsubstituted or substituted by one or more OH, N(R6)2, COOH, halogen, )C=O or XC1-5alkyl;
R10 is R3 or R4;
R11 is Ar, C3-8cycloalkyl, C1-8alkyl, C2-8alkenyl, C2-8alkynyl all of which may be unsubstituted or substituted by one or more OH, CH2OH, N(R6)2 or halogen;
R12 is hydrogen, C1-6alkyl, C2-6alkenyl or C2-7alkynyl;
X is (CH2)n, O, NR6 or S(O)q;
Y is CH3 or X(CH2)nAr,
Ar is:
Figure imgf000026_0001
Figure imgf000026_0002
naphthyl, indolyl, pyridyl, thienyl, oxazolidinyl, oxazolyl, thiazolyl, isothiazolyl, pyrazolyl, triazolyl, tetrazolyl, imidazolyl, imidazolidinyl,
thiazolidinyl, isoxazolyl, oxadiazolyl, thiadiazolyl, morpholinyl, piperidinyl, piperazinyl, pyrrolyl, or pyrimidyl; all of which may be unsubstituted or substituted by one or more R3 or R4 groups;
A is C=O, or [C(R6)2]m;
B is -CH2- or -O-;
Z1 and Z2 are independently hydrogen, C1-8alkyl, C2-8alkenyl, C2- 8alkynyl, OH, C1-8alkoxy, S(O)qC1-8alkyl, N(R6)2, Br, F, I, Cl, NHCOR6, -X(CH2)nR8, phenyl, benzyl or C3-6cycloalkyl wherein the C1-8alkyl, C2- 8alkenyl or C2-8alkynyl may be optionally substituted by COOH, OH,
CO(CH2)nCH3, CO(CH2)nCH2N(R6)2, or halogen; or Z1 and Z2 together may be -O-A-O- on contiguous carbons;
Z3 is Z1 orXR9Y;
q is zero, one or two;
n is an integer from 0 to six; m is 1, 2 or 3; and the dotted line indicates the optional presence of one or two double bonds; or a pharmaceutically acceptable salt thereof; provided that
. R2 is not hydrogen when X is S(O)q;
. when an optional double bond is present tiiere is only one R1o and when adjacent to P1 and P2, there is no P1 and P2 is not NR6R9Y;
. when an optional double bond is present in Formula (I) and X-R2 is attached to the double bond, X is not NR6;
. when an optional double bond is present and R1 is attached directly to the double bond, R1 is not NR6Ar;
. when R3, R5, Z1, Z2, or Z3 is X(CH2)nR8 and n is O, X is not oxygen or NR6 when R8 is OR6 or CO2H; and
. when R8 is CO(CR11)2O(CO)XR7, X is not S(O)q; and
. when there are two double bonds present and R1and R2 are phenyl; R3, R4 and R5 are not all hydrogen.
2. A compound of claim 1 wherein R1 is X(CH2)nAr, (Ar is (a) or (b)), dihydrobenzofuranyl, benzodioxanyl, cyclohexyl, or C1-4alkyl; R2 is (a), (b), C1-4alkyl, indolyl or hydrogen; R3 and R5 are independently hydrogen, OH, C1-5alkoxy, halogen, -OC1 alkyl phenyl, R11CO2R7, C1-4alkyl, N(R6)2, NH(CO)CH3, -X(CH2)nR8, -XR9 pyridyl, phenyl or S(O)p C1-5alkyl; R4 is hydrogen, OH, C1-5alkoxy, halogen, C1-4alkyl, N(R6)2, NH(CO)CH3 or
S(O)pC1-5alkyl; Z1, Z2 and Z3 are independently XR9Y, benzyl, hydrogen, OH, C1-5alkoxy, - N(R6)2, S(O)qC1-8alkyl, NHCOR6, X(CH2)nR8 or halogen, or Z1 and Z2 together may be -O-A-O on contiguous carbons; P1 and P2 are
independentiy hydrogen, CO2H or tetrazole; Ar is (a), (b), phenyl, or pyridyl; X is (CH2)n or oxygen.
3. A compound of claim 2 wherein R3 is hydrogen or -X(CH2)nR8, R11CO2R7; R4 and R5 are independentiy hydrogen, OH, C1-5alkoxy, SC1-5alkyl, F, Br, C1-3alkyl or NH2; Z1 and Z3 are hydrogen and Z2 is hydrogen, OH, C1-5alkoxy, halogen, X(CH2)nR8, NH2, benzyl, NH(CO)CH3, or Z1 and Z2 togetiier may be O-A-O.
4. A compound of claim 3 wherein R1 is (a) or (b); R2 is (a) or (b); A is
CH2, B is -O-; there is no optional double bond; R1 and XR2 are trans to P1; Z2 is OH, C1-5alkoxy, -OCH2CHCH2 or hydrogen, Z1 and Z3 are hydrogen; R3 is XAr, hydrogen, X(CH2)qCOOH, X(CH2)qCONR7SO2R11 or CH=CHCO2H, R4 is hydrogen, substituted phenyl, or C1-2alkoxy; and R5, R10 and P2 are hydrogen.
5.. A pharmaceutical composition comprising a compound of claim 1 and a pharmaceutically acceptable carrier.
6. A method of antagonizing endothelin receptors which comprises administering to a subject in need thereof, an effective amount to antagonize endothelin receptors or a compound of claim 1.
7. A method of treating hypertension, renal failure or cerebrovascular disease which comprises administering to a subject in need thereof, an effective amount of a compound of claim 1.
PCT/US1994/008917 1993-08-06 1994-08-03 Endothelin receptor antagonists Ceased WO1995004530A1 (en)

Priority Applications (2)

Application Number Priority Date Filing Date Title
EP94925208A EP0711157A4 (en) 1993-08-06 1994-08-03 Endothelin receptor antagonists
JP7506557A JPH09501178A (en) 1993-08-06 1994-08-03 Endothelin receptor antagonist

Applications Claiming Priority (4)

Application Number Priority Date Filing Date Title
US10349593A 1993-08-06 1993-08-06
US26923194A 1994-06-29 1994-06-29
US08/103,495 1994-06-29
US08/269,231 1994-06-29

Publications (1)

Publication Number Publication Date
WO1995004530A1 true WO1995004530A1 (en) 1995-02-16

Family

ID=26800519

Family Applications (1)

Application Number Title Priority Date Filing Date
PCT/US1994/008917 Ceased WO1995004530A1 (en) 1993-08-06 1994-08-03 Endothelin receptor antagonists

Country Status (4)

Country Link
US (1) US5817683A (en)
EP (1) EP0711157A4 (en)
JP (1) JPH09501178A (en)
WO (1) WO1995004530A1 (en)

Cited By (2)

* Cited by examiner, † Cited by third party
Publication number Priority date Publication date Assignee Title
WO1997025321A3 (en) * 1996-01-05 1997-09-12 Texas Biotechnology Corp Substituted phenyl compounds and derivatives thereof that modulate the activity of endothelin
US5962531A (en) * 1997-01-28 1999-10-05 Syntex (U.S.A.) Inc. 5-aroylnaphthalene derivatives as anti-inflammatory agents

Families Citing this family (1)

* Cited by examiner, † Cited by third party
Publication number Priority date Publication date Assignee Title
EP1130027A1 (en) * 2000-02-29 2001-09-05 Aventis Pharma Deutschland GmbH Memno peptides, process for preparation and use thereof

Citations (1)

* Cited by examiner, † Cited by third party
Publication number Priority date Publication date Assignee Title
US4401676A (en) * 1977-06-01 1983-08-30 Merck & Co., Inc. Novel α-amino acids

Family Cites Families (4)

* Cited by examiner, † Cited by third party
Publication number Priority date Publication date Assignee Title
US3562257A (en) * 1967-10-28 1971-02-09 Tanabe Seiyaku Co Benzothiazepine derivatives
US4963568A (en) * 1989-05-31 1990-10-16 Abbott Laboratories Dopamine agonists
IT1237526B (en) * 1989-12-06 1993-06-08 Zambon Spa INTERMEDIATE PREPARATION PROCESS FOR DILTIAZEM
PL176250B1 (en) * 1991-11-05 1999-05-31 Smithkline Beecham Corp Method of obtaining indane derivatives

Patent Citations (1)

* Cited by examiner, † Cited by third party
Publication number Priority date Publication date Assignee Title
US4401676A (en) * 1977-06-01 1983-08-30 Merck & Co., Inc. Novel α-amino acids

Cited By (4)

* Cited by examiner, † Cited by third party
Publication number Priority date Publication date Assignee Title
WO1997025321A3 (en) * 1996-01-05 1997-09-12 Texas Biotechnology Corp Substituted phenyl compounds and derivatives thereof that modulate the activity of endothelin
US5977117A (en) * 1996-01-05 1999-11-02 Texas Biotechnology Corporation Substituted phenyl compounds and derivatives thereof that modulate the activity of endothelin
US5962531A (en) * 1997-01-28 1999-10-05 Syntex (U.S.A.) Inc. 5-aroylnaphthalene derivatives as anti-inflammatory agents
US6150397A (en) * 1997-01-28 2000-11-21 Syntex (U.S.A.) Inc. 5-aroylnaphthalene derivatives as anti-inflammatory agents

Also Published As

Publication number Publication date
EP0711157A4 (en) 1996-07-31
EP0711157A1 (en) 1996-05-15
US5817683A (en) 1998-10-06
JPH09501178A (en) 1997-02-04

Similar Documents

Publication Publication Date Title
EP0650484B1 (en) Endothelin receptor antagonists
EP0699069B1 (en) Endothelin receptor antagonists
US6075037A (en) Endothelin receptor antagonists
WO1994014434A1 (en) Endothelin receptor antagonists
US5719183A (en) Endothelin receptor antagonists
AU3125993A (en) Endothelin receptor antagonists
US5834469A (en) Endothelin receptor antagonists
US5686481A (en) Endothelin receptor antagonists
US5559105A (en) Endothelin receptor antagonists
US5736564A (en) Endothelin receptor antagonists
US5817653A (en) Endothelin receptor antagonists
US5817683A (en) Endothelin receptor antagonists
US5929106A (en) Endothelin receptor antagonists
US6620826B2 (en) Endothelin receptor antagonists
US5716985A (en) Endothelin receptor antagonists
EP0693051B1 (en) Cyclopentene and cyclopentane derivatives as endothelin receptor antagonists
WO1995013817A1 (en) Endothelin converting enzyme inhibitors
US5929116A (en) Cyclopentene derivatives as endothelin receptor antagonists
WO1996030358A1 (en) Phenyl derivatives useful as endothelin receptor antagonists
HK1012252B (en) Endothelin receptor antagonists

Legal Events

Date Code Title Description
AK Designated states

Kind code of ref document: A1

Designated state(s): JP US

AL Designated countries for regional patents

Kind code of ref document: A1

Designated state(s): AT BE CH DE DK ES FR GB GR IE IT LU MC NL PT SE

121 Ep: the epo has been informed by wipo that ep was designated in this application
DFPE Request for preliminary examination filed prior to expiration of 19th month from priority date (pct application filed before 20040101)
WWE Wipo information: entry into national phase

Ref document number: 1994925208

Country of ref document: EP

WWE Wipo information: entry into national phase

Ref document number: 08591516

Country of ref document: US

WWP Wipo information: published in national office

Ref document number: 1994925208

Country of ref document: EP

WWW Wipo information: withdrawn in national office

Ref document number: 1994925208

Country of ref document: EP