WO1995018124A1 - Substituted morpholine derivatives and their use as therapeutic agents - Google Patents
Substituted morpholine derivatives and their use as therapeutic agents Download PDFInfo
- Publication number
- WO1995018124A1 WO1995018124A1 PCT/GB1994/002819 GB9402819W WO9518124A1 WO 1995018124 A1 WO1995018124 A1 WO 1995018124A1 GB 9402819 W GB9402819 W GB 9402819W WO 9518124 A1 WO9518124 A1 WO 9518124A1
- Authority
- WO
- WIPO (PCT)
- Prior art keywords
- phenyl
- compound
- bis
- trifluoromethyl
- formula
- Prior art date
- Legal status (The legal status is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the status listed.)
- Ceased
Links
- 0 CC(CCC1CC1)C(C)C1C(*)=CC2(CC2)C1 Chemical compound CC(CCC1CC1)C(C)C1C(*)=CC2(CC2)C1 0.000 description 2
- YWKSINPSASCIMZ-UHFFFAOYSA-N CC1NC=NC1C Chemical compound CC1NC=NC1C YWKSINPSASCIMZ-UHFFFAOYSA-N 0.000 description 1
- VALUMXGSLBMNES-UHFFFAOYSA-N Cc1n[nH]nc1C Chemical compound Cc1n[nH]nc1C VALUMXGSLBMNES-UHFFFAOYSA-N 0.000 description 1
Classifications
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D413/00—Heterocyclic compounds containing two or more hetero rings, at least one ring having nitrogen and oxygen atoms as the only ring hetero atoms
- C07D413/02—Heterocyclic compounds containing two or more hetero rings, at least one ring having nitrogen and oxygen atoms as the only ring hetero atoms containing two hetero rings
- C07D413/06—Heterocyclic compounds containing two or more hetero rings, at least one ring having nitrogen and oxygen atoms as the only ring hetero atoms containing two hetero rings linked by a carbon chain containing only aliphatic carbon atoms
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P1/00—Drugs for disorders of the alimentary tract or the digestive system
- A61P1/08—Drugs for disorders of the alimentary tract or the digestive system for nausea, cinetosis or vertigo; Antiemetics
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P11/00—Drugs for disorders of the respiratory system
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P25/00—Drugs for disorders of the nervous system
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P25/00—Drugs for disorders of the nervous system
- A61P25/04—Centrally acting analgesics, e.g. opioids
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P25/00—Drugs for disorders of the nervous system
- A61P25/06—Antimigraine agents
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P25/00—Drugs for disorders of the nervous system
- A61P25/28—Drugs for disorders of the nervous system for treating neurodegenerative disorders of the central nervous system, e.g. nootropic agents, cognition enhancers, drugs for treating Alzheimer's disease or other forms of dementia
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P29/00—Non-central analgesic, antipyretic or antiinflammatory agents, e.g. antirheumatic agents; Non-steroidal antiinflammatory drugs [NSAID]
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P37/00—Drugs for immunological or allergic disorders
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P43/00—Drugs for specific purposes, not provided for in groups A61P1/00-A61P41/00
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D413/00—Heterocyclic compounds containing two or more hetero rings, at least one ring having nitrogen and oxygen atoms as the only ring hetero atoms
- C07D413/14—Heterocyclic compounds containing two or more hetero rings, at least one ring having nitrogen and oxygen atoms as the only ring hetero atoms containing three or more hetero rings
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07F—ACYCLIC, CARBOCYCLIC OR HETEROCYCLIC COMPOUNDS CONTAINING ELEMENTS OTHER THAN CARBON, HYDROGEN, HALOGEN, OXYGEN, NITROGEN, SULFUR, SELENIUM OR TELLURIUM
- C07F9/00—Compounds containing elements of Groups 5 or 15 of the Periodic Table
- C07F9/02—Phosphorus compounds
- C07F9/547—Heterocyclic compounds, e.g. containing phosphorus as a ring hetero atom
- C07F9/6527—Heterocyclic compounds, e.g. containing phosphorus as a ring hetero atom having nitrogen and oxygen atoms as the only ring hetero atoms
- C07F9/6533—Six-membered rings
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07F—ACYCLIC, CARBOCYCLIC OR HETEROCYCLIC COMPOUNDS CONTAINING ELEMENTS OTHER THAN CARBON, HYDROGEN, HALOGEN, OXYGEN, NITROGEN, SULFUR, SELENIUM OR TELLURIUM
- C07F9/00—Compounds containing elements of Groups 5 or 15 of the Periodic Table
- C07F9/02—Phosphorus compounds
- C07F9/547—Heterocyclic compounds, e.g. containing phosphorus as a ring hetero atom
- C07F9/6558—Heterocyclic compounds, e.g. containing phosphorus as a ring hetero atom containing at least two different or differently substituted hetero rings neither condensed among themselves nor condensed with a common carbocyclic ring or ring system
- C07F9/65583—Heterocyclic compounds, e.g. containing phosphorus as a ring hetero atom containing at least two different or differently substituted hetero rings neither condensed among themselves nor condensed with a common carbocyclic ring or ring system each of the hetero rings containing nitrogen as ring hetero atom
Definitions
- This invention relates to a class of aromatic compounds which are useful as tachykinin antagonists. More particularly, the compounds of the invention contain an amine-substituted azo-heterocyclic moiety.
- tachykinins are a group of naturally occurring peptides found widely distributed throughout mammalian tissues, both within the
- the tachykinins are distinguished by a conserved carboxyl-terminal sequence: Phe-X-Gly-Leu-Met-NH 2
- neurokinin A (NKA, substance K, neuromedin L) and neurokinin B (NKB, neuromedin K)
- NKA neurokinin A
- NKB neuromedin K
- the current nomenclature designates the three tachykinin receptors mediating the biological actions of substance P, NKA and NKB as the UK., NK_ and NK_ receptors, respectively.
- substance P is believed inter alia to be involved in the neurotransmission of pain sensations [Otsuka et al, "Role of Substance P as a Sensory Transmitter in Spinal Cord and Sympathetic Ganglia” in 1982 Substance P in the Nervous System, Ciba Foundation Symposium 91 , 13-34 (published by Pitman) and Otsuka and Yanagisawa, "Does Substance P Act as a Pain Transmitter?" TIPS (1987) 8, 506-510], specifically in the transmission of pain in migraine (B.E.B. Sandberg et al, J. Med Chem, (1982) 25, 1009) and in arthritis [Levine et al Science (1984) 226, 547-549].
- Tachykinins have also been ' implicated in gastrointestinal (Gl) disorders and diseases of the Gl tract such as inflammatory bowel disease [Mantyh et al Neuroscience (1988) 25(3), 817-37 and D. Regoli in “Trends in Cluster Headache” Ed. Sicuteri et al Elsevier Scientific Publishers, Amsterdam (1987) page 85)] and emesis [F. D. Tattersall et al, Eur. J. Pharmacol., (1993) 250, R5-R6].
- substance P is believed to be involved in the inflammatory response in diseases such as rheumatoid arthritis and osteoarthritis, and fibrositis [O'Byrne et al, Arthritis and Rheumatism (1990) 33, 1023-8].
- diseases such as rheumatoid arthritis and osteoarthritis, and fibrositis [O'Byrne et al, Arthritis and Rheumatism (1990) 33, 1023-8].
- Other disease areas where tachykinin antagonists are believed to be useful are allergic conditions [Hamelet et al, Can. J. Pharmacol.
- Tachykinin antagonists may also be useful in the treatment of 5 small cell carcinomas, in particular small cell lung cancer (SCLC) [Langdon et al, Cancer Research (1992) 52, 4554-7].
- SCLC small cell lung cancer
- Substance P may also play a role in demyelinating diseases such as multiple sclerosis and amyotrophic lateral sclerosis [J. Luber-Narod et al, poster C.I.N.P. XVIIIth Congress, 28th June-2nd July 1992], and in io disorders of bladder function such as bladder detrusor hyper-reflexia ⁇ Lancet, 16th May 1992, 1239).
- demyelinating diseases such as multiple sclerosis and amyotrophic lateral sclerosis [J. Luber-Narod et al, poster C.I.N.P. XVIIIth Congress, 28th June-2nd July 1992]
- io disorders of bladder function such as bladder detrusor hyper-reflexia ⁇ Lancet, 16th May 1992, 1239).
- tachykinins have utility in the following disorders: depression, dysthymic disorders, chronic obstructive airways disease, hypersensitivity disorders such as poison ivy, 15 vasospastic diseases such as angina and Reynauld's disease, fibrosing and collagen diseases such as scleroderma and eosinophilic fascioliasis, reflex sympathetic dystrophy such as shoulder/hand syndrome, addiction disorders such as alcoholism, stress related somatic disorders, neuropathy, neuralgia, disorders related to immune enhancement or suppression such as systemic 20 lupus erythmatosus (European patent specification no.
- R is a large variety of substituents
- R and R are inter alia hydrogen
- R is inter alia
- R is inter alia optionally substituted phenyl
- R , R and R are a variety of substituents
- X a is O, S, SO or SO 2 ;
- Y a is inter alia O
- Z a is hydrogen or C ⁇ alkyl.
- the compounds of the present invention possess a particularly advantageous profile of activity having potent antagonist activity at the NK T receptor and a long duration of action.
- the compounds of the present invention, and in particular their pharmaceutically acceptable acid addition salts, are also particularly suited to a wide variety of pharmaceutical formulations by virtue of their stability.
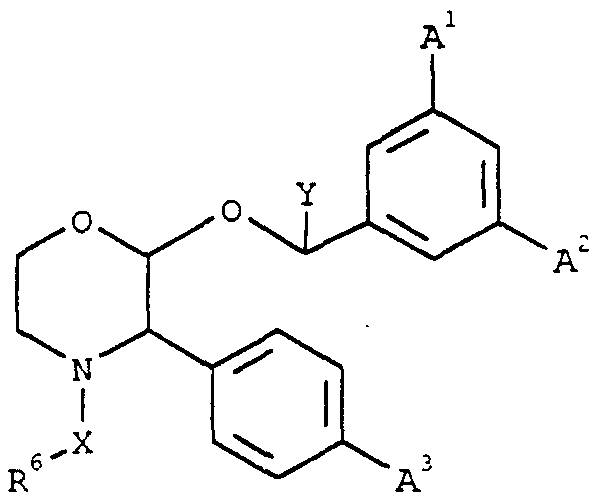
- the present invention provides compounds of the formula (I):
- R is hydrogen, halogen, C. alkyl, C 1-6 alkoxy, CF NO,, CN, a b
- R is hydrogen, halogen, C alkyl, C alkoxy substituted by
- R is hydrogen, halogen or CF 3 ;
- R is hydrogen, halogen, C ⁇ alkyl, C ⁇ alkoxy, CF 3 , NO 2 , CN, SR a , SOR a , S0 2 R a , C0 2 R a , CONR a R b , C ⁇ alkenyl, C M alkynyl or C ⁇ alkyl a b substituted by C ⁇ alkoxy, where R and R each independently represent hydrogen or C 1 _ 4 alkyl; R 5 is hydrogen, halogen, C 1 _ 6 alkyl, C ⁇ alkoxy substituted by
- R 7 is hydrogen, C 1 _ 4 alkyl, C ⁇ 7 cycloalkyl or C 3 7 cycloalkylC 1 4 alkyl, or C 2 4 alkyl substituted by C. 4 al oxy or hydroxyl; 8
- R is hydrogen, C ⁇ alkyl, O-.-.cycloalkyl or C g .-.cycloalkylC ⁇ alkyl, or C ⁇ alkyl substituted by one or two substituents selected from C ⁇ alkoxy, hydroxyl or a 4, 5 or 6 membered heteroaliphatic ring containing one or two heteroatoms selected from N, O and S;
- R 7 8 or R , R and the nitrogen atom to which they are attached form a heteroaliphatic ring of 4 to 7 ring atoms, optionally substituted by one or two groups selected from hydroxy or optionally substituted by a or hydroxyl group, and optionally containing a double bond, which ring may optionally contain an oxygen or sulphur ring atom, a group S(O) or S(O) 2 or a second nitrogen atom which will be part of a NH or NR moiety where R is C ⁇ alkyl optionally substituted by hydroxy or C 1 l alkoxy; or R 7 , R 8 and the nitrogen atom to which they are attached form a non-aromatic azabicyclic ring system of 6 to 12 ring atoms; or Z, R and the nitrogen atom to which they are attached form a heteroaliphatic ring of 4 to 7 ring atoms which may optionally contain an oxygen ring atom;
- R and R are each independently hydrogen or C. ⁇ alkyl, or R
- Y is a C 1 4 alkyl group optionally substituted by a hydroxyl group; with the proviso that if Y is C ⁇ . 4 alkyl, R 6 is substituted at least by a group of formula ZNR 7 R 8 as defined above; and pharmaceutically acceptable salts and prodrugs thereof.
- R is hydrogen, C. ⁇ alkyl, C 1 4 alkoxy, halo or CF 3 .
- R is hydrogen, C. ⁇ alkyl, C ⁇ alkoxy, halogen or CF 3 .
- R is hydrogen, fluorine, chlorine or CF 3 .
- Favourably R is fluorine, chlorine or CF 3 .
- R is hydrogen, fluorine, chlorine or CF 3 .
- R is hydrogen, fluorine, chforine or CF 3 .
- R and R are in the 3 and 5 positions of the phenyl ring.
- R is 3-fluoro or 3-CF3.
- R is 5-fluoro or 5-CF3.
- R is hydrogen
- R is 3-F or 3-CF.
- R is 5-CF, and R is hydrogen.
- R is hydrogen
- R is hydrogen, fluorine, chlorine or CF 3 .
- R is hydrogen and R is hydrogen or 4-fluoro.
- R and R are each independently hydrogen or methyl.
- R is hydrogen.
- R is hydrogen.
- R and R are both hydrogen.
- A is fluorine or CF 3 ;
- 2 A is fluorine or CF,
- A is fluorine or hydrogen
- a preferred class of compound of formula (I) or (la) is that wherein Y represents a group substituted by a hydroxy group; or a pharmaceutically acceptable salt or prodrug thereof.
- another preferred class of compound of formula (I) or (la) is that wherein Y represents a C h alky! group, with the proviso that R 6 is substituted at least by a group of the formula ZNR 7 R 8 as defined above; or a pharmaceutically acceptable salt or prodrug thereof.
- R is hydrogen, C ⁇ alkyl, C ⁇ cycloalkyl or C ⁇ 7 cycloalkylC 1-4 alkyl, or C ⁇ alkyl substituted by one or two substituents selected from C. ⁇ alkoxy, hydroxyl or a 4, 5 or 6 membered heteroaliphatic ring containing one or two heteroatoms selected from N, O and S;
- R 7 8 or R , R and the nitrogen atom to which they are attached form a heteroaliphatic ring of 4 to 7 ring atoms, optionally substituted by a hydroxy group, and optionally containing a double bond, which ring may optionally contain an oxygen or sulphur ring atom, a group S(O) or S(O) 2 or a second nitrogen atom which will be part of a NH or NR moiety where R is C ⁇ alkyl optionally substituted by hydroxy or C ⁇ alkoxy; or Z, R and the nitrogen atom to which they are attached form a heteroaliphatic ring of 4 to 7 ring atoms which may optionally contain an oxygen ring atom; or a pharmaceutically acceptable salt or prodrug thereof.
- a preferred class of compounds of formula (I) or (la) is that wherein
- Y represents a group
- Z is C ⁇ alkylene or C ⁇ cycloalkylene
- R is hydrogen or C ⁇ alkyl, or C ⁇ alkyl substituted by C ⁇ alkoxy
- R is hydrogen or C ⁇ alkyl or C ⁇ alkyl substituted by C. ⁇ alkoxy, hydroxyl or a 5 or 6 membered heteroaliphatic ring containing one or two heteroatoms selected from N, O and S;
- R 7 8 or R , R and the nitrogen atom to which they are attached form a heteroaliphatic ring of 4 to 7 ring atoms, optionally substituted by a hydroxy group, which ring may optionally contain an oxygen or sulphur ring atom, a group S(O) or S(O) 2 or a second nitrogen atom which will be part of a NH or NR C moiety where R c is C. ⁇ alkyl optionally substituted by hydroxy or C ⁇ alkoxy;
- R and the nitrogen atom to which they are attached form a heteroaliphatic ring of 4 to 7 ring atoms which may optionally contain an oxygen ring atom; or a pharmaceutically acceptable salt or prodrug thereof.
- another preferred class of compound of formula (I) or (la) is that wherein Y represents a d ⁇ alkyl group substituted by a hydroxyl group;
- Z is C ⁇ alkylene or C ⁇ cycloalkylene
- R is hydrogen or C 1 ⁇ alkyl, or C ⁇ alkyl substituted by C ⁇ alkoxy
- R is hydrogen or C 1 J alkyl or C ⁇ alkyl substituted by C. ⁇ alkoxy or hydroxyl;
- R 7 8 or R , R and the nitrogen atom to which they are attached form a heteroaliphatic ring of 4 to 7 ring atoms which may optionally contain an oxygen ring atom or a second nitrogen atom which will be part of a NH or NR C moiety where R c is C ⁇ alkyl optionally substituted by hydroxy or C ⁇ alkoxy; or Z, R and the nitrogen atom to which they are attached form a heteroaliphatic ring of 4 to 7 ring atoms which may optionally contain an oxygen ring atom; or a pharmaceutically acceptable salt or prodrug thereof.
- Z is C 1 6 alkylene or C 3 ⁇ cycloalkyl
- R is hydrogen or C 1 4 alkyl, or C. ⁇ alkyl substituted by C. ⁇ alkoxy
- R is hydrogen or C 1 4 alkyl or C 2 4 alkyl substituted by C ⁇ alkoxy or hydroxyl;
- R 7 8 or R , R and the nitrogen atom to which they are attached form a heteroaliphatic ring of 4 to 7 ring atoms which may optionally contain an oxygen ring atom or a second nitrogen atom which will be part of a NH or NR moiety where R is C ⁇ alkyl optionally substituted by hydroxy or C ⁇ alkoxy; or Z, R and the nitrogen atom to which that are attached form a heteroaliphatic ring to 4 to 7 ring atoms which may optionally contain an oxygen ring atom; or a pharmaceutically acceptable salt or prodrug thereof.
- a preferred group Y for compounds of the formulae (I) or (la) is the CH 2 OH group.
- Another preferred group Y for compounds of the formulae (I) or (la) is the CH 3 group.
- Particularly apt values for X for compounds of the formulae (I) or (la) include CH 2 , CH(CH 3 ) and CH 2 CH 2 of which the CH 2 group is preferred.
- R is a 5-membered ring
- R Particularly preferred heterocyclic rings represented by R 6 are selected from:
- R may represent a heterocyclic ring selected from:
- a particularly preferred heterocyclic ring represented by R is:
- a further favoured group of compounds of the present invention are of the formula (lc) and pharmaceutically acceptable salts and prodrugs thereof:
- a 1 , A 2 and A 3 are as defined in relation to formula (la) and Q 1 is CH or N or C-ZNR 7 R 8 wherein Z, R 7 and R 8 are as defined in relation to formula
- Z may be a linear, branched or cyclic group.
- Favourably Z contains 1 to 4 carbon atoms and most favourably 1 or 2 carbon atoms.
- a particularly favourable group Z is CH 2 .
- R may aptly be a C ⁇ alkyl group or a C. ⁇ alkyl group substituted by
- R may aptly be a C 1 ⁇ alkyl group or a C ⁇ alkyl
- R 7 8 group substituted by a hydroxyl or C 1 2 alkoxy group or R and R may be linked so that, together with the nitrogen atom to which they are attached, they form an azetidinyl, pyrrolidinyl, piperidyl, morpholino, thiomorpholino, piperazino or piperazino group substituted on the nitrogen atom by a C ⁇ alkyl group or a C 2 ⁇ alkyl group substituted by a hydroxy or C 1 2 alkoxy group.
- group NR R represents a heteroaliphatic ring of 4 to
- a particularly preferred group is 3-pyrroline.
- group NR 7 R 8 represents a non-aromatic azabicyclic ring system
- such a system may contain between 6 and 12, and preferably between 7 and 10, ring atoms.
- Suitable rings include
- R represents a C- ⁇ alkyl group substituted by a 5 or 6 membered heteroaliphatic ring containing one or two heteroatoms selected from N, 0 and S
- suitable rings include pyrrolidino, piperidino, piperazino, morpholino, or thiomorpholino.
- Particularly preferred are nitrogen containing heteroaliphatic rings, especially pyrrolidino and morpholino rings. 7 8
- moieties ZNR R include those wherein Z
- NR R is amino, methyiamino, dimethylamino, diethylamino, azetidinyl, pyrrolidino and morpholino.
- ZNR R represents hydrogen, C ⁇ alkyl
- R is C h alky I substituted by one or two substituents selected from hydroxy, C 1 2 alkoxy, azetidinyl, pyrrolidino, piperidino, morpholino or thiomorpholino.
- Z is preferably CH 2 and NR 7 R 8 is preferably dimethylamino, azetidinyl or pyrrolidino, especially dimethylamino.
- A is preferably fluorine or CF 3 ; A is preferably CF 3 ; and A is preferably fluorine.
- alkyl or "alkoxy" as a group or part of a group means that the group is straight or branched.
- suitable alkyl groups include methyl, ethyl, n-propyl, i-propyl, n-butyl, s-butyl and t-butyl.
- suitable alkoxy groups include methoxy, ethoxy, n-propoxy, i-propoxy, n-butoxy, s-butoxy and t-butoxy.
- cycloalkyl groups referred to herein may represent, for example, cyclopropyl, cyclobutyl, cyclopentyl or cyclohexyl.
- a suitable cycloalkylalkyl group may be, for example, cyclopropylmethyl.
- alkenyl and “alkynyl” as a group or part of a group means that the group is straight or branched.
- suitable alkenyl groups include vinyl and allyl.
- a suitable alkynyl group is propargyl.
- halogen means fluorine, chlorine, bromine and iodine.
- the most apt halogens are fluorine and chlorine of which fluorine is preferred.
- Specific compounds within the scope of this invention include: 2-(R)-(1-(R)-(3,5-bis(trifluoromethyl)phenyl)ethoxy)-4-(2,3-dihydro-5-(N,N- dimethylamino)methyl-2-oxo-1 ,3-imidazol-4-yl)methyl-3-(S)-(4-fluorophenyl) morpholine;
- Still further preferred compounds within the scope of the present invention include: 2-(R)-(1-(S)-(3,5-bis(trifluoromethyl)phenyl)-2-hydroxyethoxy)-3-(S)-(4- fluorophenyl)-4-(2,3-dihydro-3-oxo-1 ,2,4-triazol-5-yl)methylmorpholine;
- the compounds of formula (I) will preferably be prepared in the form of a pharmaceutically acceptable salt, especially an acid addition salt.
- the salts of the compounds of formula (I) will be non-toxic pharmaceutically acceptable salts.
- Other salts may, however, be useful in the preparation of the compounds according to the invention or of their non-toxic pharmaceutically acceptable salts.
- Suitable pharmaceutically acceptable salts of the compounds of this invention include acid addition salts which may, for example, be formed by mixing a solution of the compound according to the invention with a solution of a pharmaceutically acceptable acid such as hydrochloric acid, fumaric acid, p-toluenesulphonic acid, maleic acid, succinic acid, acetic acid, citric acid, tartaric acid, carbonic acid, phosphoric acid or sulphuric acid.
- a pharmaceutically acceptable acid such as hydrochloric acid, fumaric acid, p-toluenesulphonic acid, maleic acid, succinic acid, acetic acid, citric acid, tartaric acid, carbonic acid, phosphoric acid or sulphuric acid.
- Salts of amine groups may also comprise quaternary ammonium salts in which the amino nitrogen atom carries a suitable organic group such as an alkyl, alkenyl, alkynyl or aralkyl moiety.
- suitable pharmaceutically acceptable salts thereof may include metal salts such as alkali metal salts, e.g. sodium or potassium salts; and alkaline earth metal salts, e.g. calcium or magnesium salts.
- prodrugs of the compounds of formula (I) above include within its scope prodrugs of the compounds of formula (I) above.
- prodrugs will be functional derivatives of the compounds of formula (I) which are readily convertible in vivo into the required compound of formula (I).
- Conventional procedures for the selection and preparation of suitable prodrug derivatives are described, for example, in "Design of Prodrugs", ed. H. Bundgaard, Elsevier, 1985.
- a prodrug may be a pharmacologically inactive derivative of a biologically active substance (the "parent drug” or "parent molecule") that requires transformation within the body in order to release the active drug, and that has improved delivery properties over the parent drug molecule.
- the transformation in vivo may be, for example, as the result of some metabolic process, such as chemical or enzymatic hydrolysis of a carboxylic, phosphoric or sulphate ester, or reduction or oxidation of a susceptible functionality.
- certain preferred prodrugs may not be antagonists of tachykinin, particularly substance P, activity to any significant extent (or not at all).
- Such compounds are still advantageous in treating the various conditions described herein, especially where an injectable formulation is preferred.
- a prodrug may lie in its physical properties, such as enhanced water solubility for parenteral administration compared with the parent drug, or it may enhance absorption from the digestive tract, or it may enhance drug stability for long-term storage.
- a prodrug will improve the overall efficacy of a parent drug, for example, through the reduction of toxicity and unwanted effects of drugs by controlling their absorption, blood levels, metabolism, distribution and cellular uptake.
- a particularly preferred class of prodrugs of the compounds of the present invention is that wherein the hydroxyl moiety of the group Y in formula (I) (when Y if Chalky! substituted by hydroxyl) is derivatized.
- Suitable prodrug derivatives include:
- + M is a pharmaceutically acceptable monovalent counterion
- D is a pharmaceutically acceptable divalent counterion
- R is hydrogen or C 1 3 alkyl; and R is a group selected from -O(CH 2 ) 2 NH 3 + .M “ ; -O(CH, 2)',2NH, 2(R 12 ) / + . • M • " " ;> -OCH- ',2CO W 2 " ' M + ;
- M is a pharmaceutically acceptable monovalent counterion
- R is hydrogen, C 1 4 alkyl or C 2 ⁇ alkyl substituted by a hydroxyl or C. 4 alkoxy group.
- prodrug derivatives are: (b) -(CHR 1 °) n -PO(O-) 2 .2M + ;
- parent molecule refers to the biologically active entity that is released via enzymatic action of a metabolic or catabolic process, or via a chemical process following administration of the prodrug.
- the parent compound may also be the starting material for the preparation of its corresponding prodrug.
- the preferred routes of administration are oral and intravenous.
- the prodrugs are hydrolyzed or otherwise cleaved in vivo to the corresponding parent compounds of formula (I), or a pharmaceutically acceptable salt thereof. Since the parent compounds may less than optimally soluble in aqueous solutions, the above prodrugs provide a distinct advantage by virtue of their relatively enhanced aqueous solubility.
- Examples of negative monovalent counterions defined herein as "M" include acetate, adipate, benzoate, benzenesulfonate, bisulfate, butyrate, camphorate, camphorsulfonate, citrate, ethanesulfonate, fumarate, hemisulfate, 2-hydroxyethylsulfonate, heptanoate, hexanoate, hydrochloride, hydrobromide, hydroiodide, lactate, malate, maleate, methanesulfonate, 2-naphthalenesulfonate, oxalate, pamoate, persulfate, picrate, pivalate, propionate, salicylate, stearate, succinate, sulfate, tartrate, tosylate (p-toluenesulfonate), and undecanoate.
- Base salts which are pharmaceutically acceptable monovalent cations defined herein as "M" or pharmaceutically acceptable divalent
- 2+ cations defined herein as "D" include ammonium salts, alkali metal salts such as sodium, lithium and potassium salts, alkaline earth metal salts such as aluminium, calcium and magnesium salts, salts with organic bases such as dicyclohexylamine salts, N-methyl-D-glucamine, and salts with
- each of M may be the same or different.
- a divalent cation D may instead be present.
- the basic nitrogen-containing groups may be quatemized with such agents as: lower alkyl halides, such as methyl, ethyl, propyl, and butyl chlorides, bromides and iodides; dialkyl sulfates like dimethyl, diethyl and dibutyl; diamyl sulfates; long chain halides such as decyl, lauryl, myristyl and stearyl chlorides, bromides and iodides; aralkyl halides like benzyl bromide and others.
- the non-toxic physiologically acceptable salts are preferred, although other salts are also useful, such as in isolating or purifying the product.
- the salts may be formed by conventional means, such as by reacting the free base form of the product with one or more equivalents of the appropriate acid in a solvent or medium in which the salt is insoluble, or in a solvent such as water which is removed in vacuo or by freeze drying or by exchanging the anions of an existing salt for another anion on a suitable ion exchange resin.
- a particularly preferred sub-class of prodrugs of the compounds of the present invention is that defined by the formula (le) and pharmaceutically acceptable salts thereof:
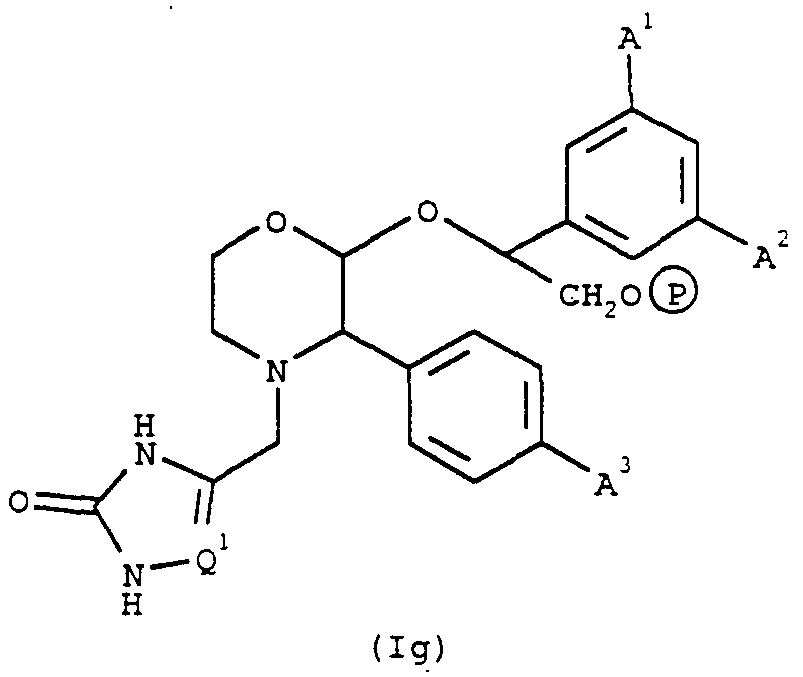
- R , R , R , R , R , R and X are as defined in relation to formula (I) and P in a circle is PO(OH)0 " .M + , PO(0 " ) 2M + , or PO(0"),.D 2+ .
- Another preferred sub-class of prodrugs of the compounds of the present invention is that defined by the formula (If) and pharmaceutically acceptable salts thereof:
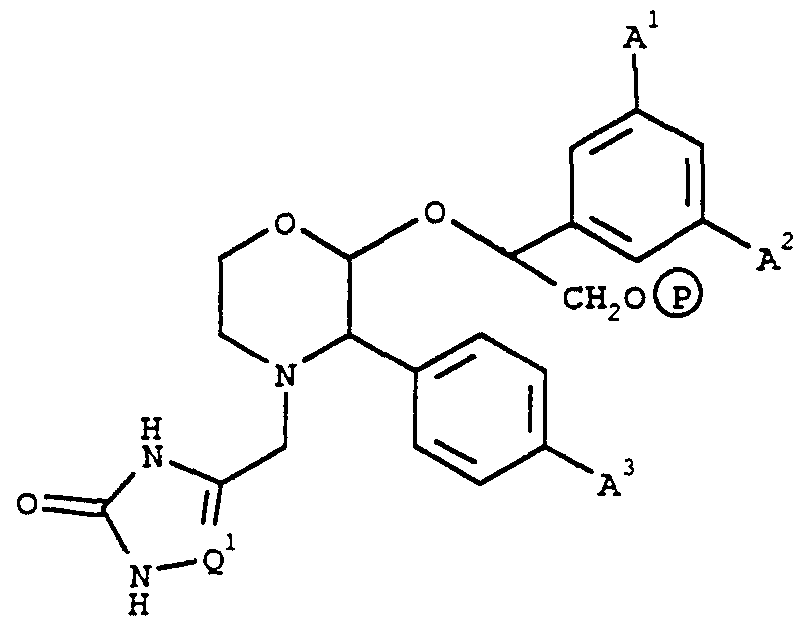
- An especially preferred sub-group of prodrugs of the compounds of the present invention is that defined by formula (Ig) and pharmaceutically acceptable salts thereof:
- a yet further preferred sub-group of prodrugs of the compounds of the present invention is that defined by formula (Ih) and pharmaceutically acceptable salts thereof:
- prodrug derivatives within the scope of this invention include: 2-(R)-(1-(S)-(3,5-bis(trifluoromethyl)phenyl)-2-phosphoryloxyethoxy)-3-(S)-(4- fluorophenyl)-4-(2,3-dihydro-3-oxo-1 ,2,4-triazol-5-yl)methylmorpholine;
- 2 3 is preferably fluorine or CF 3 ;
- A is preferably CF 3 ; and
- A is preferably fluorine.
- the present invention includes within its scope solvates of the compounds of formula (I) and salts thereof, for example, hydrates.
- the compounds according to the invention have at least three asymmetric centres, and may accordingly exist both as enantiomers and as diastereoisomers. It is to be understood that all such isomers and mixtures thereof are encompassed within the scope of the present invention.
- the preferred compounds of the formula (I), (la), (lb), (lc), (Id), (le), (If), (Ig) and (Ih) will have the 2- and 3- substituent cis and the preferred stereochemistry at the 2-position is that possessed by the compound of Example 1 (i.e. 2-(R)-), the preferred stereochemistry of the 3-position is that possessed by the compound of Example 1 (i.e. 3-(S)), and the preferred stereochemistry of the carbon to which the group Y is either (R) when Y is Ci ialkyl (e.g. methyl) or (S) when Y is C h alky! substituted by hydroxy (e.g. CH 2 OH).
- the present invention further provides pharmaceutical compositions comprising one or more compounds of formula (I) in association with a pharmaceutically acceptable carrier.
- compositions according to the invention are in unit dosage forms such as tablets, pills, capsules, powders, granules, solutions or suspensions, or suppositories, for oral, parenteral or rectal administration, or administration by inhalation or insufflation.
- a pharmaceutical carrier e.g. conventional tableting ingredients such as corn starch, lactose, sucrose, sorbitol, talc, stearic acid, magnesium stearate, dicalcium phosphate or gums, and other pharmaceutical diluents, e.g.
- a solid preformulation composition containing a homogeneous mixture of a compound of the present invention, or a non-toxic pharmaceutically acceptable salt thereof.
- preformulation compositions as homogeneous, it is meant that the active ingredient is dispersed evenly throughout the composition so that the composition may be readily subdivided into equally effective unit dosage forms such as tablets, pills and capsules.
- This solid preformulation composition is then subdivided into unit dosage forms of the type described above containing from 0.1 to about 500 mg of the active ingredient of the present invention.
- the tablets or pills of the novel composition can be coated or otherwise compounded to provide a dosage form affording the advantage of prolonged action.
- the tablet or pill can comprise an inner dosage and an outer dosage component, the latter being in the form of an envelope over the former.
- the two components can be separated by an enteric layer which serves to resist disintegration in the stomach and permits the inner component to pass intact into the duodenum or to be delayed in release.
- enteric layers or coatings such materials including a number of polymeric acids and mixtures of polymeric acids with such materials as shellac, cetyl alcohol and cellulose acetate.
- liquid forms in which the novel compositions of the present invention may be incorporated for administration orally or by injection include aqueous solutions, suitably flavoured syrups, aqueous or oil suspensions, and flavoured emulsions with edible oils such as cottonseed oil, sesame oil, coconut oil or peanut oil, as well as elixirs and similar pharmaceutical vehicles.
- Suitable dispersing or suspending agents for aqueous suspensions include synthetic and natural gums such as tragacanth, acacia, alginate, dextran, sodium carboxymethylcellulose, methylcellulose, polyvinyl- pyrrolidone or gelatin.
- compositions for administration by injection include those comprising a compound of formula (I), as the active ingredient, in association with a surface-active agent (or wetting agent or surfactant) or in the form of an emulsion (as a water-in-oil or oil-in-water emulsion).
- a surface-active agent or wetting agent or surfactant
- an emulsion as a water-in-oil or oil-in-water emulsion
- Suitable surface-active agents include anionic agents such as sodium bis-(2-ethylhexyl)sulfosuccinate (docusate sodium), cationic agents, such as alkyltrimethylammonium bromides, (e.g. cetyltrimethylammonium bromide (cetrimide)), and in particular, non-ionic agents, such as polyoxyethylenesorbitans (e.g. TweenTM 20, 40, 60, 80 or 85) and other sorbitans (e.g. SpanTM 20, 40, 60, 80 or 85).
- Compositions with a surface-active agent will conveniently comprise between 0.05 and 5% surface-active agent, and preferably between 0.1 and 2.5%. It will be appreciated that other ingredients may be added, for example mannitol or other pharmaceutically acceptable vehicles, if necessary.
- Suitable emulsions may be prepared using commercially available fat emulsions, such as IntralipidTM, LiposynTM, InfonutrolTM,
- LipofundinTM and LipiphysanTM may be either dissolved in a pre-mixed emulsion composition or alternatively it may be dissolved in an oil (e.g. soybean oil, safflower oil, cottonseed oil, sesame oil, corn oil or almond oil) and an emulsion formed upon mixing with a phospholipid (e.g. egg phospholipids, soybean phospholipids or soybean lecithin) and water.
- a phospholipid e.g. egg phospholipids, soybean phospholipids or soybean lecithin
- Suitable emulsions will typically contain up to 20% oil, for example, between 5 and 20%.
- the fat emulsion will preferably comprise fat droplets between 0.1 and 1.0 ⁇ m, particularly 0.1 and 0.5 ⁇ m, and have a pH in the range of 5.5 to 8.0.
- Particularly preferred emulsion compositions are those prepared by mixing a compound of formula (I) with IntralipidTM or the components thereof (soybean oil, egg phospholipids, glycerol and water).
- compositions for inhalation or insufflation include solutions and suspensions in pharmaceutically acceptable, aqueous or organic solvents, or mixtures thereof, and powders.
- the liquid or solid compositions may contain suitable pharmaceutically acceptable excipients as set out above.
- the compositions are administered by the oral or nasal respiratory route for local or systemic effect.
- Compositions in preferably sterile pharmaceutically acceptable solvents may be nebulised by use of inert gases. Nebulised solutions may be breathed directly from the nebulising device or the nebulising device may be attached to a face mask, tent or intermittent positive pressure breathing machine.
- Solution, suspension or powder compositions may be administered, preferably orally or nasally, from devices which deliver the formulation in an appropriate manner.
- the present invention futher provides a process for the preparation of a pharmaceutical composition comprising a compound of formula (I), which process comprises bringing a compound of formula (I) into association with a pharmaceutically acceptable carrier or excipient.
- the compounds of formula (I) are of value in the treatment of a wide variety of clinical conditions which are characterised by the presence of an excess of tachykinin, in particular substance P, activity.
- disorders of the central nervous system such as anxiety, depression, psychosis and schizophrenia; epilepsy; neurodegenerative disorders such as dementia, including senile dementia of the Alzheimer type, Alzheimer's disease and Down's syndrome; demyelinating diseases such as MS and ALS and other neuropathological disorders such as peripheral neuropathy, for example diabetic and chemotherapy-induced neuropathy, and postherpetic and other neuralgias; small cell carcinomas such as small cell lung cancer; respiratory diseases, particularly those associated with excess mucus secretion such as chronic obstructive airways disease, bronchopneumonia, chronic bronchitis, cystic fibrosis and asthma, and bronchospasm; inflammatory diseases such as inflammatory bowel disease, psoriasis, fibrositis, osteoarthritis, rheumatoid
- the compounds of formula (I) are also of value in the treatment of a combination of the above conditions, in particular in the treatment of combined post-operative pain and post-operative nausea and vomiting.
- the compounds of formula (I) are particularly useful in the treatment of emesis, including acute, delayed or anticipatory emesis, such as emesis induced by chemotherapy, radiation, toxins, pregnancy, vestibular disorders, motion, surgery, migraine, and variations in intercranial pressure.
- the compounds of formula (I) are of use in the treatment of emesis induced by antineoplastic (cytotoxic) agents including those routinely used in cancer chemotherapy.
- chemotherapeutic agents include alkylating agents, for example, nitrogen mustards, ethyleneimine compounds, alkyl sulphonates and other compounds with an alkylating action such as nitrosoureas, cisplatin and dacarbazine; antimetabolites, for example, folic acid, purine or pyrimidine antagonists; mitotic inhibitors, for example, vinca alkaloids and derivatives of podophyllotoxin; and cytotoxic antibiotics.
- alkylating agents for example, nitrogen mustards, ethyleneimine compounds, alkyl sulphonates and other compounds with an alkylating action such as nitrosoureas, cisplatin and dacarbazine
- antimetabolites for example, folic acid, purine or pyrimidine antagonists
- mitotic inhibitors for example, vinca alkaloids and derivatives of podophyllotoxin
- cytotoxic antibiotics include cytotoxic antibiotics.
- chemotherapeutic agents are described, for instance, by D. J. Stewart in Nausea and Vomiting: Recent Research and Clinical Advances, Eds. J. Kucharczyk et al, CRC Press Inc., Boca Raton, Florida, USA (1991 ) pages 177-203, especially page 188.
- Commonly used chemotherapeutic agents include cisplatin, dacarbazine (DTIC), dactinomycin, mechlorethamine (nitrogen mustard), streptozocin, cyclophosphamide, carmustine (BCNU), lomustine (CCNU), doxorubicin
- the compounds of formula (I) are also of use in the treatment of emesis induced by radiation including radiation therapy such as in the treatment of cancer, or radiation sickness; and in the treatment of post ⁇ operative nausea and vomiting.
- the compounds of formula (I) may be presented together with another therapeutic agent as a combined preparation for simultaneous, separate or sequential use for the relief of emesis. Such combined preparations may be, for example, in the form of a twin pack.
- a further aspect of the present invention comprises the compounds of formula (I) in combination with a 5-HT 3 antagonist, such as ondansetron, granisetron or tropisetron, or other anti-emetic medicaments, for example, a dopamine antagonist such as metoclopramide.
- a compound of formula (I) may be administered in combination with an anti- inflammatory corticosteroid, such as dexamethasone.
- a compound of formula (I) may be administered in combination with a chemotherapeutic agent such as an alkylating agent, antimetabolite, mitotic inhibitor or cytotoxic antibiotic, as described above.
- a chemotherapeutic agent such as an alkylating agent, antimetabolite, mitotic inhibitor or cytotoxic antibiotic, as described above.
- the currently available dosage forms of the known therapeutic agents for use in such combinations will be suitable.
- the compounds of the present invention were found to attenuate the retching and vomiting induced by cisplatin.
- the compounds of formula (I) are also particularly useful in the treatment of pain or nociception and/or inflammation and disorders associated therewith such as, for example, neuropathy, such as diabetic and chemotherapy-induced neuropathy, postherpetic and other neuralgias, asthma, osteroarthritis, rheumatoid arthritis and especially migraine.
- neuropathy such as diabetic and chemotherapy-induced neuropathy, postherpetic and other neuralgias, asthma, osteroarthritis, rheumatoid arthritis and especially migraine.
- the present invention further provides a compound of formula (I) for use in therapy.
- the present invention provides a compound of formula (I) for use in the manufacture of a medicament for the treatment of physiological disorders associated with an excess of tachykinins, especially substance P.
- the present invention also provides a method for the the treatment or prevention of physiological disorders associated with an excess of tachykinins, especially substance P, which method comprises administration to a patient in need thereof of a tachykinin reducing amount of a compound of formula (I) or a composition comprising a compound of formula (I).
- a compound according to the present invention for the treatment of certain conditions it may be desirable to employ a compound according to the present invention in conjunction with another pharmacologically active agent.
- a compound of formula (I) may be used in conjunction with a bronchodilator, such as a ⁇ 2-adrenergic receptor antagonist or tachykinin antagonist which acts at NK-2 receptors.
- the compound of formula (I) and the bronchodilator may be administered to a patient simultaneously, sequentially or in combination.
- the present invention accordingly provides a method for the treatment of a respiratory disease, such as asthma, which method comprises administration to a patient in need thereof of an effective amount of a compound of formula (I) and an effective amount of a bronchodilator.
- a respiratory disease such as asthma
- the present invention also provides a composition comprising a compound of formula (I), a bronchodilator, and a pharmaceutically acceptable carrier.
- the excellent pharmacological profile of the compounds of the present invention offers the opportunity for their use in therapy at low doses thereby minimising the risk of unwanted side effects.
- a suitable dosage level is about 0.001 to 50 mg/kg per day, in particular about 0.01 to about 25 mg/kg, such as from about 0.05 to about 10 mg/kg per day.
- a suitable dosage level is about 0.001 to 25 mg/kg per day, preferably about 0.005 to 10 mg/kg per day, and especially about 0.005 to 5 mg/kg per day.
- the compounds may be administered on a regimen of 1 to 4 times per day, preferably once or twice per day.
- a suitable dosage level is about 0.001 to 10 mg/kg per day, preferably about 0.005 to 5 mg/kg per day, and especially 0.01 to 2 mg/kg per day.
- the compounds may be administered on a regimen of 1 to 4 times per day, preferably once or twice per day.
- the compounds according to the invention may be prepared from compounds of formula (II)
- G 7 ⁇ wherein R represents 1 ,2,3-thazol-4-yl substituted by CH 2 NR R , and X is -CH -, may be prepared by reaction of a compound of formula (IV) with an azide, for example, sodium azide in a suitable solvent such as dimethylsulphoxide at a temperature of between 40°C and 100°C, followed
- R represents substituted or unsubstituted 1 ,3,5-triazine
- reaction is conveniently effected in a suitable organic solvent, such as acetonitrile, at elevated temperature, such as 80-90°C, preferably about 82°C.
- a suitable organic solvent such as acetonitrile
- R represents substituted or unsubstituted 1 ,2,4-triazine
- reaction is conveniently effected in a suitable organic solvent, such as an ether, e.g. tetrahydrofuran, conveniently at ambient temperature.
- a suitable organic solvent such as an ether, e.g. tetrahydrofuran, conveniently at ambient temperature.
- R represents a substituted 1 ,2,4-triazolyl group
- R is H, CONH 2 or OCH 3 (which is converted to an oxo substituent under the reaction conditions), in the presence of a base, followed where necessary by conversion to a compound of formula (I), for example, by reduction of the CONH 2 group to CH 2 NH 2 .
- Suitable bases of use in the reaction include alkali metal carbonates such as, for example, potassium carbonate.
- the reaction is conveniently effected in an anhydrous organic solvent such as, for example, anhydrous dimethylformamide, preferably at elevated temperature, such as about 140°C.
- a suitable reducing agent for the group CONH 2 is lithium aluminium hydride, used at between -10°C and room temperature. According to another process, (G), compounds of formula (I)
- R represents thioxotriazolyl
- Suitable bases of use in the reaction include organic bases such as, for example, 1 ,8-diazabicyclo[5.4.0]undec-7-ene (DBU).
- DBU 1 ,8-diazabicyclo[5.4.0]undec-7-ene
- the reaction is conveniently effected in a suitable organic solvent, such as alcohol, e.g. butanol.
- Compounds of formula (I) may also be prepared from other compounds of formula (I) using suitable interconversion procedures.
- compounds of formula (I) wherein X represents C alkyl may be prepared from compounds of formula (I) wherein X represents C 1 4 alkyl substituted by oxo by reduction, for example, using borane or lithium aluminium hydride.
- Suitable interconversion procedures will be readily apparent to those skilled in the art.
- Intermediates of formula (IV) may be prepared from intermediates of formula (II) by reaction with an acetylene compound of formula HC ⁇ C-CH 2 -Hal in the presence of a base such as potassium carbonate in a suitable solvent such as dimethylformamide, conveniently at room temperature, followed by reaction of the resultant acetylene
- Hal is a halogen atom, for example, chlorine, bromine or iodine, especially chlorine, by reaction with an azide, for example, sodium azide in a suitable solvent such as dimethylsulphoxide at or below room temperature.
- an azide for example, sodium azide in a suitable solvent such as dimethylsulphoxide at or below room temperature.
- Compounds of formula (XI) may be prepared by a dropwise addition of an intermediate of formula (II) to a dihaloacetylene of formula
- Hal-CH 2 -C ⁇ C-CH 2 -Hal where each Hal is independently chlorine, bromine or iodine, especially chlorine.
- the reaction is conveniently effected in a suitable solvent such as dimethylformamide in the presence of a base such as potassium carbonate.
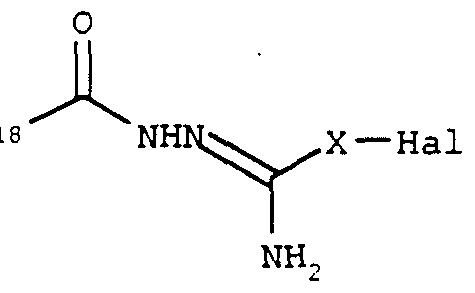
- Intermediates of formula (VI) may be prepared from intermediates of formula (II) by reaction with a compound of formula Hal-X-C(NH)NH 2 , where Hal and X are as previously defined.
- Intermediates of formula (VII) may be prepared from intermediates of formula (II) by reaction with a compound of formula Hal-X-C(NH)NHNH-Boc, wherein Hal and X are as previously defined and Boc stands for t-butoxycarbonyl, followed by deprotection under acidic conditions.
- Compounds of formula (VIII) are commercially available or may be prepared from commercially available compounds by known methods.
- imidazolinone group lacking a CH 2 NR R may be reacted with paraformaldehyde and an amine for example a secondary amine such as pyrrolidine to give a compound wherein the imidazolinone ring is substituted
- R , R and the nitrogen atom to which they are attached form a heteroaliphatic ring of 4 to 7 ring atoms which may optionally contain an oxygen ring atom or a second nitrogen atom which will be part of a NH or NR moiety, where R is as previously defined.
- a further alternative method for the preparation of certain compounds of formula (I) involves the reaction of an intermediate of formula (II) as defined above with one of the compounds of formula (XII):
- This reaction is conveniently effected in an organic solvent such as dimethylformamide in the presence of an acid acceptor such as potassium carbonate.
- an acid acceptor such as potassium carbonate.
- reactive groups may be protected, thus for example, the NH groups of an imidazolinone of formula (Xlla) may be protected by any suitable amine protecting group such as an acetyl group.
- the preferred phosphate prodrugs of the compounds of the present invention may be prepared in a stepwise manner from a compound of formula (I) wherein Y is, for example, -CH 2 OH-.
- the hydroxy compound is first treated with dibenzyloxydiethylaminophosphine in a suitable solvent such as tetrahydrofuran, preferably in the presence of an acid catalyst such as tetrazole.
- a suitable solvent such as tetrahydrofuran
- an acid catalyst such as tetrazole.
- the hydroxy compound of formula (I) may be reacted with a suitable base such as sodium hydride in tetrahydrofuran, and tetrabenzylpyrophosphate added to yield the dibenzyl- protected phosphate which may be deprotected as described above.
- a suitable base such as sodium hydride in tetrahydrofuran
- tetrabenzylpyrophosphate added to yield the dibenzyl- protected phosphate which may be deprotected as described above.
- L-Selectride is lithium tri-sec-butylborohydride.
- the protecting groups may be removed at a convenient subsequent stage using methods known from the art.
- the exemplified compounds of this invention were tested by the methods set out at pages 36 to 39 of International Patent Specification No.
- WO 93/01 165 The compounds or, in the case of prodrugs, the parent compounds, were found to be active with IC ⁇ at the NKi receptor of less than
- Step A 3-(4-Fluorophenyl)acetyl-4-(S)-benzyl-2-oxazolidinone
- the solution was cooled to -10°C and treated with 5.60ml (40.0mmol) of t ethylamine followed by 4.30ml (35.0mmol) of trimethylacetyl chloride. A white precipitate formed immediately.
- the resulting mixture was stirred at -10°C for 40 minutes, then cooled to -78°C.
- Step B 3-((S)-Azido-(4-fluorophenvn)acetyl-4-(S)-benzyl-2- oxazolidinone
- Step A' (4-Fluorophenyl)acetyl chloride
- a solution of 150g (0.974mol) of 4-(fluorophenyl)acetic acid and 1 ml of N.N-dimethylformamide in 500ml of toluene at 40°C was treated with 20ml of thionyl chloride and heated to 40°C.
- An additional 61.2ml of thionyl chloride was added dropwise over 1.5 hours.
- Step C Methyl ( ⁇ H4-fluorophenyl)glvcine
- a solution of 24.7g (0.1 mol) of methyl 2-bromo-2-(4- fluorophenyl)acetate (from Step B') and 2.28g (0.01 mol) of benzyl triethylammonium chloride in 25ml of methanol was treated with 6.8g (0.105mol) of sodium azide and the resulting mixture was stirred for 20 hours at room temperature.
- the reaction mixture was filtered; the filtrate was diluted with 50ml of methanol and hydrogenated in the presence of 0.5g of 10% Pd/C at 50 psi for 1 hour.
- the solution was filtered and the solvent removed in vacuo.
- the two crops of the (S)-amino ester (39.8g) were combined in 200ml of 7:1 v/v ethanol/water, heated for 30 minutes and cooled to room temperature. Addition of ethyl acetate, cooling, and filtration afforded 31.7g of (S)-(4- fluorophenyl)glycinate, (+)-DBT salt (ee > 98%). Enantiomeric excess was determined by chiral HPLC (Crownpak CR(+) 5% MeOH in aq HCIO 4 pH2 1.5ml/min 40°C 200nm).
- the reaction mixture was concentrated in vacuo and the residue was dissolved in 40ml of water.
- the aqueous solution was washed (3 x 30ml of ethyl acetate) and the layers were separated.
- Step A N-Benzyl-(SH4-fluorophenv ⁇ lvcine
- a solution of 1.87g (11.05mmol) of (S)-(4-fluorophenyl)-glycine (from Description 1 ) and 1.12ml (11.1 mmol) of benzaldehyde in 11.1 ml of 1 N aqueous sodium hydroxide solution and 11 ml of methanol at 0°C was treated with 165mg (4.4mmol) of sodium borohydride. The cooling bath was removed and the resulting mixture was stirred at room temperature for 30 minutes.
- Step B 4-Benzyl-3-(SM4-fluorophenvO-2-morpholinone A mixture of 1.95g (7.5mmol) of N-benzyl (S)-(4- fluorophenyl)glycine, 3.90ml (22.5mmol) of N,N-diisopropyl- ethylamine, 6.50ml (75.0mmol) of 1 ,2-dibromoethane and 40ml of N,N-dimethylformamide was stirred at 100°C for 20 hours
- the dimethyl titanocene could be stored as a solution in toluene at 0°C for at least 2 weeks without apparent chemical degradation.
- 1 H NMR 200MHz, CDCI 3 ) ⁇ -0.15 (6H, s), 6.06 (1 OH, s).
- Step B 4-Benzyl-2-(R)-(1-(3.5-bis(trifluoromethvnphenvn ethenyloxy)-3-(SM4-fluorophenv0morpholine
- a solution of the compound of Description 3 (2.50g, 4.9mmol) and 2.50g (12.0mmol) of dimethyl titanocene (from Step A) in 35ml of 1:1 v/v THF/toluene was stirred in an oil bath at 80°C for 16 hours. The reaction mixture was cooled and concentrated in vacuo.
- Step B 4-Benzyl-3-(S)-Phenyl-2-morpholinone A mixture of 4.00g (16.6mmol) of N-benzyl-(S)-phenylglycine (from Step A) 5.00g (36.0mmol) of potassium carbonate, 10.0ml of 1 ,2-dibromoethane and 25ml of N,N-dimethylformamide was stirred at 100°C for 20 hours. The mixture was cooled and partitioned between 200ml of ethyl ether and 100ml of water. The layers were separated and the organic layer was washed with 3 x 50ml of water, dried over magnesium sulfate and concentrated in vacuo.
- DESCRIPTION 17 4-Benzyl-2-(RH1 -(3.5-bis(trifluoromethvnphenyl) ethenyloxy)-3- (S)-phenylmorpholine
- a solution of 2.50g (4.9mmol) of the compound of Description 16 and 2.50g (12.0mmol) of dimethyl titanocene (Description 4a), in 35ml of 1 :1 v/v THF/toluene was stirred in an oil bath at 80°C for 16 hours. The reaction mixture was cooled and concentrated in vacuo.
- the compound of Description 20 (8g) was dissolved in ethyl acetate (100ml) and isopropanol (50ml) and palladium on charcoal (1.5g) was added to the solution. This mixture was hydrogenated at 40 psi overnight. The catalyst was removed by filtration and the solvents were removed in vacuo. The residue was purified by flash silica chromatography using 1-10% methanol in dichloromethane as eluant. This afforded the product as a white powder (5.7g, 90%).
- Step A 4-Benzyl-3- SH4-fluorophenvn-2-(RH1-(SH3-fluoro-5-
- Step B 3-(SH4-Fluorophenyl)-2-(R 1-(SH3-fluoro-5- (thfluoromethyl)phenyl)-2-hvdroxyethoxy)morpholine
- the compound of (a) above (1g) was dissolved in ethyl acetate/2-propanol (20ml, 3:1 ) and treated with Pd on carbon (100mg). The mixture was hydrogenated at 60 psi for 12h. The catalyst was removed by filtration and the solvent was removed in vacuo. The residue was purified by medium pressure chromatography on silica (Lobar) using 5% methanol in dichloromethane as eluant. The product was recrystallised from ether.
- Step A 4-Benzyl-2-(R)-(1-fS)-(3.5-Bis(trifluoromethvnphenvn-2- hvdroxyethoxy)-3-(S)-phenylmorpholine
- the compound of Description 17 was reacted with diborane and subsequently with basic hydrogen peroxide according to the method illustrated in Description 20. This intermediate was not purified and was reacted crude in the following step.
- Step B 2-(RH1-(SH3.5-Bis(trifluoromethyl)phenyl-2- hvdroxyethoxy)-3-fS)-phenylmorpholine
- Step A 4-Benzyl-2-mH1-(SH3-fluoro-5-ttrifluoromethvn phenyl)-2-hvdroxyethoxy)-3-(S)-phenylmorpholine
- Step B 2- R)-(1-(S)-(3-fluoro-5-(trifluoromethyl)phenyl)-2- hydro ⁇ yethoxy)-3-(S)-phenylmorpholine
- Step A 4-Benzyl-3-(S)-phenyl-2-(R 1-(SH3- (t ⁇ fluoromethyl)phenyl)-2-hydroxyethoxy)morpholine Prepared from the compound of Description 29 following the method illustrated in Description 20.
- Step B 3-fS)-Phenyl-2-(R)-f1-.S)-f3-(trifluoromethyl)phenylV2- hvdroxyethoxy)morpholine This was carried through without purification to the title compound following the method illustrated in Description 21.
- reaction mixture was stirred for 10 min and, pyrrol idine (1.1g) was added in one portion and stirring was continued for 20 min.
- the reaction mixture was diluted with water (250ml) and extracted with ethyl acetate (3 x 50ml). The combined organic extracts were washed with water (2 x 50ml) and brine (1 x 50ml) and then dried (K2CO3) and concentrated in vacuo.
- the residue was purified by chromatography on silica using a gradient elution of dichloromethane (100%) to dichloromethane/methanol/aqueous ammonia mixtures (85:15:0.5) to provide the title compound as a foam.
- Propargyl bromide (1.9ml) was added to a stirred mixture of the compound of Description 5 (5g) and potassium carbonate (4.76g) in dry dimethylformamide at 23°C. After 15 min the reaction mixture was diluted with water (250ml) and extracted with ethyl acetate (3 x 100ml). The combined organic phases were washed with brine (1 x 100ml) then dried (K 2 CO 3 ) and concentrated to leave an oil. This was purified by chromatography on silica using ethyl acetate in hexane (1 :9 then 1 :4) as eluent to afford the title compound as an oil.
- Lithium aluminium hydride (0.47ml, 1 M in tetrahydrofuran) was added dropwise to a solution of the compound described in (c) above (0.11g) in dry tetrahydrofuran (1ml) under an inert atmosphere at 23°C. After 30 min sodium hydroxide (10 drops, 1M) was added followed by water (5 drops). Ethyl acetate (50ml) was then added and the resulting mixture was filtered through a pad of Hyflo. The filtrate was concentrated in vacuo and the residue was purified by chromatography on silica using ethyl acetate in methanol (9:1 then 4:1 ) as eluant to provide the title compound as a foam.
- Dimethylamine (approximately 10ml) was condensed at -80°C in a pressure tube and to this was added a solution of N-(4-azidobut- 2-ynyl)-2-(R)-(1-(R)-(3,5-bis(trifluoromethyl)phenyl) ethoxy)-3-(S)- (4-fluorophenyl)morpholine (3.2g) in dioxan (15ml). The tube was sealed and the solution was heated at 90°C for 16h. The solution was evaporated to dryness and the residue chromatographed on silica gel (eluting with 5% methanol in dichloromethane containing 0.25% ammonia (SG.
- silica gel eluting with 5% methanol in dichloromethane containing 0.25% ammonia
- Examples 14 to 21 in Table 2 were prepared in a similar manner to that described in Example 12, Method B, via the appropriate N-(4-azidobut-2-ynyl)morpholine and the appropriate amine.
- EXAMPLE 22 2-(R)- ⁇ -(R)-f3.5-Bis ⁇ rifluoromethvnphenyl)ethoxy)-3-(S)- phenyl-4-(1-(2-pyrrolidinoethyl)-1 ,2.4-triazol-3-yl)methylmorpholine
- Lithium aluminium hydride (1.0M solution in tetrahydrofuran, 1.9ml) was added to a solution of the compound described in (a) above (isomer B) in tetrahydrofuran (5ml) at 0°C. The mixture was warmed to room temperature and was stirred for 1h. The mixture was quenched (sodium hydroxide and water) and filtered through celite to remove inorganics. The filtrate was evaporated and purified on silica using 10% methanol in dichloromethane as eluent. This afforded the product as a yellow oil.
- This compound was prepared by the method described in Example 12 (Method A) and purified by chromatography on silica using ethyl acetate, petroleum ether (60-80°C) and methanol (3:10:0, then 1 :0:0 followed by 9:0:1 ) as eluent to afford the title compound as a white foam.
- Examples 25 to 27 in Table 2 were prepared in a similar manner to that described in Example 12, Method B, via the appropriate N-(4-azidobut-2-ynyl)morpholine and the appropriate amine.
- Example 7 The product from Example 7 (0.2g) and phosphorus oxychloride (0.5ml) was heated at reflux for 20 hours. The mixture was cooled and partitioned between dichloromethane and aqueous potassium carbonate solution. The organic layer was washed (H 2 O), dried (MgSO 4 ) and evaporated in vacuo. The product was purified by chromatography on silica using 100% ethyl acetate followed by 5% methanol:95% ethyl acetate to afford the title compound as an oil.
- Examples 31 to 37 in Table 2 were prepared in a similar manner to that described in Example 12, Method B, via the appropriate N-(4-azidobut-2-ynyl)morpholine and the appropriate amine.
- Examples 38 to 41 in Table 1 were prepared in a similar manner to that described in Example 4 from the appropriate morpholine, 4,5-bis(bromomethyl)-1 ,3-diacetyl-2-imidazolinone and the appropriate amine.
- Examples 43 to 62 in Table 2 were prepared in a similar manner to that described in Example 12, Method B, via the appropriate N-(4-azidobut-2-ynyl)morpholine and the appropriate amine.
- Examples 64 to 74 in Table 3 were prepared in a similar manner to that described in Example 63 from the appropriate 1 ,2,4-triazol-3- ylmethylmorpholine and the appropriate amine.
- EXAMPLE 75 2-(R)-(1-(R)-3.5-Bisftrifluoromethvnphenyl)ethoxy)-4-(5-(N.N- dimethylaminomethvO-1 ,2.4-triazol-3-vOmethyl-3-(S)-(4- fluorophenvPmorpholine
- EXAMPLE 76 4-(5-(N,N-Dimethylaminomethyl)-1.2.3-triazol-4-yl)methyl-3-fS)- (4-fluorophenyl)-2-(R)-(1-(R)-(3-methylthio-5-(trifluoromethvn phenyl)ethoxy)morpholine
- the compound of Example 57 (270mg, 0.51 mmol) was heated to 120°C with sodium thiomethoxide (178mg, 2.55mmol) in anhydrous DMF (10ml) for between 2-5 hours.
- Example 102 4-(5-(N,N-Dimethylaminomethv ⁇ -1.2.3-triazol-4-yl)methyl-2-(R)- (1-(R)-f3-methylthio-5-(trifluoromethvnphenvnethoxy)-3-(S)- phenylmorpholine
- the title compound was prepared from the triazole of Example 102 according to the method of Example 76 as a foam (116mg, 36%).
- the title compound was prepared from the compound of Example 57 according to the method of Example 76 as a foam
- Example 76 The thioether of Example 76 (155mg, 0.28mmol) was dissolved in trifluoroacetic acid (800 ⁇ l) cooled to 0°C and treated with a 2.0M solution of trifluoroperacetic acid in trifluoroacetic acid (153 ⁇ l, 0.308mmol), with stirring for 30 minutes. The reaction mixture was poured into 0.5M sodium bicarbonate solution (50ml), extracted with dichloromethane (3 x 15ml), dried (MgSO 4 ) and concentrated in vacuo.
- Example 77 The title compound as an unresolved mixture of stereoisomers was prepared from Example 77 according to the method of Example 81 as a foam (90mg, 63%).
- Example 78 The title compound as an unresolved mixture of stereoisomers was prepared from Example 78 according to the method of Example 81 as a foam (113mg, 92%).
- 1 H NMR (360MHz,CDCI 3 ) ⁇ 1.3 and 1.41 (3H total, 2 x d, J 6.6), 2.54 and 2.57 (3H total, 2 x s), 2.54- 2.65 (1 H, m), 2.82-2.89 (1 H, m), 3.05-3.25 (4H, vbr s), 3.35 (1 H, m),
- EXAMPLE 84 3-(S)-(4-FluorophenylV2-(RH1- RH3-methylsulphonyl-5- (trifluoromethyl) phenyl)ethoxy)-4-(5-morpholinomethyl-1 ,2,3- triazol-4-yl)methylmorpholine
- the sulphoxide of Example 83 (78mg, 0.128mmol) was dissolved in trifluoroacetic acid (500 ⁇ l) cooled to 0°C and treated with a 2.0M solution of t fluoroperacetic acid in trifluoroacetic acid (70 ⁇ l, 0.140mmol) with stirring for 2 1 _ hours.
- a further equivalent of trifiuoroperacetic acid 70 ⁇ l, 0.140mmol was added after this time and the product purified after 3 hours according to the method of
- Example 81 to yield the title compound as a foam (27mg, 34%).
- Step B 2-(R)-(1-(R)-(3.5-Bis(trifluoromethvnphenvnethoxy)-3- (SM4-fluorophenyl -(4-hvdroxybut-3-vnvDmorpholine
- Step B The alcohol of Step B (0.42g; 1eq) was dissolved in dry THF (5ml) under N 2 and triphosgene (84mg; 0.35 eq) was added followed by pyridine (128 ⁇ l; 2.0 eq). The reaction mixture was stirred at room temperature for V ⁇ , then diluted with EtOAc and washed with H O and brine, dried (MgS0 4 ) and concentrated to leave a yellow oil. This was purified by chromatography (hexanes/EtOAc 9:1 ⁇ :1 ) to provide the title compound as a clear, viscous oil. MS m/z 538, 540 (MH + ).
- Step D N-(4-Azidobut-3-vnvn-2-(R)-(1-(R)-(3,5- bis(trifluoromethv ⁇ phenyl)ethoxy)-3-(SW4-fluorophenv ⁇ morpholine
- Step E 2-(R)-f1-fR)-(3.5-Bis(trifluoromethvnphenyl)ethoxy)-3- (S)-(4-fluorophenvn-4-(2-(5-((S)-f+)-2- methoxymethylpyrrolidinomethyl)-1.2,3-triazol-4-yl)ethyl)morpholine
- a solution of the azide of Step D (0.205g; 1 eq) and (S)-(+)-2- methoxymethylpyrrolidine (114 ⁇ l; 3 eq) was heated at 80°C under N 2 the solvent was removed in vacuo and the residue purified by chromatography using CH 2 CI 2 /MeOH/NH 3 (98:2:0.1 then 97:3:0.1 ) as eluant to provide the title compound as a white foam.
- Step A 2- -(1-fSK3.5-Bis(trifluoromethvnphenvn-2-.etf- butyldimethylsilyloxyethoxy)-3-(S)-(4-fluorophenyl)morpholine
- the product from Description 21 (2g) was dissolved in anhydrous dichloromethane (16ml), under nitrogen, and cooled to
- Step B 2-(R)-(1-(S)-(3.5-Bis(trifluoromethvnphenvn-2-terr- butyldimethylsilyloxyethoxy)-3-(SH4-fluorophenyl)-4-(4-chlorobut- 2-vnyl)morpholine
- Step C 2-(RW1-(SH3.5-Bisftrifluoromethvnphenvn-2-ter.- butyldimethylsilyloxyethoxy -(5-(N,N-dimethylaminomethvO-1 ,2,3- triazol-4-yl)methyl-3-(SM4-fluorophenv0morpholine Prepared in an analogous fashion to Steps (b) and (c) of
- Example 12 using the product of Step B, above, to afford the title compound.
- Example 101 The product from Example 101 (2g) was dissolved in N,N- dimethylformamide (4ml) at room temperature under nitrogen. lodomethane was added, followed by sodium hydride (60%) (14mg) and the mixture stirred for 16 hours. The reaction mixture was partitioned between ethylacetate and water and the organic layer was washed (H 2 O x 2, brine), dried (MgS0 ) and evaporated in vacuo. Purification by gravity silica chromatography eluting with
- the reaction mixture was cooled and diluted with water (100ml).
- the product was extracted into ethyl acetate (3 x 50ml) and the organic layer was washed with brine, dried (MgSO ) and evaporated in vacuo.
- the residue was purified by chromatography on silica using 7% methanol in dichloromethane as the eluant. This afforded the title compound (200mg, 60%) as a white solid.
- Example 88 The compound of Example 88 (200mg) in dry tetrahydrofuran (1 ml) was treated with dibenzyloxydiethylaminophosphine (200mg) and tetrazole (100mg). The reaction was stirred for 2 hours and then treated with a further 10Omg of dibenzyloxydiethyl aminophosphine followed after 1 hour by tetrazole (100mg). The reaction was stirred for a further 1 hour before adding 4-methylmorpholine-N-oxide (1.Og) and stirring for 16 hours. The reaction was poured into potassium carbonate solution and extracted into ethyl acetate.
- ammonium salt of the title compound was prepared from the compound of Example 96 using the methodology of Example 93.
- Examples 101 and 102 in Table 2 were prepared in a similar manner to that described in Example 12, Method B, via the appropriate N-(4-azidobut-2-ynyi)morpholine and the appropriate amine.
- the compound of formula (I), cellulose, lactose and a portion of the corn starch are mixed and granulated with 10% corn starch paste.
- the resulting granulation is sieved, dried and blended with the remainder of the corn starch and the magnesium stearate.
- the resulting granulation is then compressed into tablets containing 1.Omg, 2.0mg, 25.0mg, 26.0mg, 50.0mg and 10Omg of the active compound per tablet.
- the sodium phosphate, citric acid monohydrate and sodium chloride are dissolved in a portion of the water.
- the compound of formula (I) is dissolved or suspended in the solution and made up to volume.
- the white soft paraffin is heated until molten.
- the liquid paraffin and emulsifying wax are incorporated and stirred until dissolved.
- the compound of formula (I) is added and stirring continued until dispersed. The mixture is then cooled until solid.
- the compound of formula (I) is dissolved directly in a solution of the commercially available Tween 80TM (polyoxyethylenesorbitan monooleate) and 5% aqueous mannitol (isotonic).
- the compound of formula (I) is dissolved directly in the commercially available IntralipidTM (10 or 20%) to form an emulsion.
- the compound of formula (I) is dissolved in soybean oil. An emulsion is then formed by mixing this solution with the egg phospholipid, glycerol and water. The emulsion is then sealed in sterile vials.
Landscapes
- Chemical & Material Sciences (AREA)
- Health & Medical Sciences (AREA)
- Organic Chemistry (AREA)
- General Health & Medical Sciences (AREA)
- Life Sciences & Earth Sciences (AREA)
- Nuclear Medicine, Radiotherapy & Molecular Imaging (AREA)
- Medicinal Chemistry (AREA)
- Chemical Kinetics & Catalysis (AREA)
- Bioinformatics & Cheminformatics (AREA)
- Pharmacology & Pharmacy (AREA)
- General Chemical & Material Sciences (AREA)
- Animal Behavior & Ethology (AREA)
- Public Health (AREA)
- Veterinary Medicine (AREA)
- Engineering & Computer Science (AREA)
- Neurology (AREA)
- Neurosurgery (AREA)
- Biomedical Technology (AREA)
- Molecular Biology (AREA)
- Biochemistry (AREA)
- Pain & Pain Management (AREA)
- Hospice & Palliative Care (AREA)
- Otolaryngology (AREA)
- Rheumatology (AREA)
- Immunology (AREA)
- Psychiatry (AREA)
- Pulmonology (AREA)
- Pharmaceuticals Containing Other Organic And Inorganic Compounds (AREA)
- Heterocyclic Carbon Compounds Containing A Hetero Ring Having Nitrogen And Oxygen As The Only Ring Hetero Atoms (AREA)
- Plural Heterocyclic Compounds (AREA)
- Nitrogen Condensed Heterocyclic Rings (AREA)
- Medicines That Contain Protein Lipid Enzymes And Other Medicines (AREA)
Abstract
Description






Claims






Priority Applications (20)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| KR1019960703426A KR100286412B1 (en) | 1993-12-29 | 1994-12-23 | Substituted morpholine derivatives, preparation method thereof and pharmaceutical composition comprising the same |
| US08/663,201 US5612337A (en) | 1993-12-29 | 1994-12-23 | Substituted morpholine derivatives and their use as therapeutic agents |
| ES95904621T ES2164758T3 (en) | 1993-12-29 | 1994-12-23 | SUBSTITUTED MORPHOLINE DERIVATIVES AND ITS USE AS THERAPEUTIC AGENTS. |
| BR9408442A BR9408442A (en) | 1993-12-29 | 1994-12-23 | Compound pharmaceutical composition processes for treating or preventing physiological disorders associated with an excess of tachykinins and for preparing the compound and using the compound |
| JP7517856A JP3071467B2 (en) | 1993-12-29 | 1994-12-23 | Substituted morpholine derivatives and their use as therapeutic agents |
| RU96116158/04A RU2131426C1 (en) | 1993-12-29 | 1994-12-23 | Substituted morpholine derivatives, method of their synthesis, pharmaceutical composition based on said, method of decrease of tachikinin level in treatment or prophylaxis of physiological states associated with tachikinin excess |
| EP95904621A EP0737192B1 (en) | 1993-12-29 | 1994-12-23 | Substituted morpholine derivatives and their use as therapeutic agents |
| AU13223/95A AU685209C (en) | 1993-12-29 | 1994-12-23 | Substituted morpholine derivatives and their use as therapeutic agents |
| HU9601801A HU221139B1 (en) | 1993-12-29 | 1994-12-23 | Tachykinin antagonist substituted morpholine derivatives, preparation and use thereof |
| SK839-96A SK283070B6 (en) | 1993-12-29 | 1994-12-23 | Substituted morpholine derivatives and their use as therapeutic agents |
| CA002178219A CA2178219C (en) | 1993-12-29 | 1994-12-23 | Substituted morpholine derivatives and their use as therapeutic agents |
| DE69428913T DE69428913T2 (en) | 1993-12-29 | 1994-12-23 | SUBSTITUTED MORPHOLINE DERIVATIVES AND THEIR USE AS MEDICINAL PRODUCTS |
| NZ277839A NZ277839A (en) | 1993-12-29 | 1994-12-23 | Substituted morpholine derivatives, preparation and pharmaceutical compositions thereof |
| AT95904621T ATE207917T1 (en) | 1993-12-29 | 1994-12-23 | SUBSTITUTED MORPHOLINE DERIVATIVES AND THEIR USE AS MEDICINAL PRODUCTS |
| DK95904621T DK0737192T3 (en) | 1993-12-29 | 1994-12-23 | Substituted morpholine derivatives and their use as therapeutic agents |
| PL94315182A PL181214B1 (en) | 1993-12-29 | 1994-12-23 | Derivatives o substituted morpholine and their application as pharmaceutic agents |
| TW084102683A TW449588B (en) | 1993-12-29 | 1995-03-20 | Substituted morpholine derivatives and their use as tachykinin antagonists |
| BG100644A BG62616B1 (en) | 1993-12-29 | 1996-06-06 | Substituted morpholine derivatives and their application as therapeutical means |
| LVP-96-197A LV11687B (en) | 1993-12-29 | 1996-06-19 | Substituted morpholine derivatives and their use as therapeutic agents |
| NO962749A NO307090B1 (en) | 1993-12-29 | 1996-06-28 | Substituted morpholine derivatives and their use |
Applications Claiming Priority (8)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| GB9326480A GB9326480D0 (en) | 1993-12-29 | 1993-12-29 | Therapeutic agents |
| GB9407189A GB9407189D0 (en) | 1994-04-12 | 1994-04-12 | Therapeutic agents |
| GB9408065A GB9408065D0 (en) | 1994-04-22 | 1994-04-22 | Therapeutic agents |
| GB9416428A GB9416428D0 (en) | 1994-08-15 | 1994-08-15 | Therapeutic agents |
| GB9416428.2 | 1994-08-15 | ||
| GB9326480.2 | 1994-08-15 | ||
| GB9407189.1 | 1994-08-15 | ||
| GB9408065.2 | 1994-08-15 |
Publications (1)
| Publication Number | Publication Date |
|---|---|
| WO1995018124A1 true WO1995018124A1 (en) | 1995-07-06 |
Family
ID=27451102
Family Applications (1)
| Application Number | Title | Priority Date | Filing Date |
|---|---|---|---|
| PCT/GB1994/002819 Ceased WO1995018124A1 (en) | 1993-12-29 | 1994-12-23 | Substituted morpholine derivatives and their use as therapeutic agents |
Country Status (24)
| Country | Link |
|---|---|
| US (1) | US5612337A (en) |
| EP (3) | EP1462450B1 (en) |
| JP (2) | JP3071467B2 (en) |
| KR (1) | KR100286412B1 (en) |
| CN (1) | CN1067683C (en) |
| AT (3) | ATE207917T1 (en) |
| BG (1) | BG62616B1 (en) |
| BR (1) | BR9408442A (en) |
| CA (1) | CA2178219C (en) |
| CZ (1) | CZ288176B6 (en) |
| DE (3) | DE69434063D1 (en) |
| DK (1) | DK0737192T3 (en) |
| ES (1) | ES2164758T3 (en) |
| HU (1) | HU221139B1 (en) |
| IL (1) | IL112134A (en) |
| LV (1) | LV11687B (en) |
| NO (1) | NO307090B1 (en) |
| NZ (1) | NZ277839A (en) |
| PL (1) | PL181214B1 (en) |
| RU (1) | RU2131426C1 (en) |
| SG (1) | SG52217A1 (en) |
| SK (1) | SK283070B6 (en) |
| TW (1) | TW449588B (en) |
| WO (1) | WO1995018124A1 (en) |
Cited By (118)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| WO1995030674A1 (en) * | 1994-05-05 | 1995-11-16 | Merck Sharp & Dohme Limited | Morpholine derivatives and their use as antagonists of tachikinins |
| WO1996007649A1 (en) * | 1994-09-02 | 1996-03-14 | Merck Sharp & Dohme Limited | Morpholine derivatives and their use as therapeutic agents |
| WO1996019970A1 (en) * | 1994-12-23 | 1996-07-04 | Merck Sharp & Dohme Limited | Hemolysis prevention by surfactants and emulsions |
| WO1996029328A1 (en) * | 1995-03-18 | 1996-09-26 | Merck Sharp & Dohme Limited | Morpholine derivatives, compositions containing them and their use as therapeutic agents |
| WO1997002824A1 (en) * | 1995-07-08 | 1997-01-30 | Merck Sharp & Dohme Limited | Use of a tachykinin antagonist and a muscarinic antagonist and/or an antihistamine to treat motion sickness |
| WO1997018206A1 (en) * | 1995-11-14 | 1997-05-22 | Merck Sharp & Dohme Limited | Morpholine derivatives and their use as therapeutic agents |
| US5719147A (en) * | 1992-06-29 | 1998-02-17 | Merck & Co., Inc. | Morpholine and thiomorpholine tachykinin receptor antagonists |
| WO1998024442A1 (en) * | 1996-12-02 | 1998-06-11 | Merck Sharp & Dohme Limited | Use of nk-1 receptor antagonists for treating sexual dysfunction |
| WO1998024444A1 (en) * | 1996-12-02 | 1998-06-11 | Merck Sharp & Dohme Limited | Use of nk-1 receptor antagonists for treating substance use disorders |
| WO1998024439A1 (en) * | 1996-12-02 | 1998-06-11 | Merck Sharp & Dohme Limited | Use of nk-1 receptor antagonists for treating severe anxiety disorders |
| WO1998024447A1 (en) * | 1996-12-02 | 1998-06-11 | Merck Sharp & Dohme Limited | Use of nk-1 receptor antagonists for treating cognitive disorders |
| WO1998024443A1 (en) * | 1996-12-02 | 1998-06-11 | Merck Sharp & Dohme Limited | Use of nk-1 receptor antagonists for treating bipolar disorders |
| WO1998024440A1 (en) * | 1996-12-02 | 1998-06-11 | Merck Sharp & Dohme Limited | Use of nk-1 receptor antagonists for treating stress disorders |
| WO1998024441A1 (en) * | 1996-12-02 | 1998-06-11 | Merck Sharp & Dohme Limited | Use of nk-1 receptor antagonists for treating major depressive disorders with anxiety |
| WO1998024438A1 (en) * | 1996-12-02 | 1998-06-11 | Merck Sharp & Dohme Limited | Use of nk-1 receptor antagonists for treating major depressive disorders |
| WO1998024445A1 (en) * | 1996-12-02 | 1998-06-11 | Merck Sharp & Dohme Limited | Use of nk-1 receptor antagonists for treating schizophrenic disorders |
| WO1998024446A1 (en) * | 1996-12-02 | 1998-06-11 | Merck Sharp & Dohme Limited | Use of nk-1 receptor antagonists for treating movement disorders |
| WO1998047513A1 (en) * | 1997-04-24 | 1998-10-29 | Merck Sharp & Dohme Limited | Use of nk-1 receptor antagonists for treating eating disorders |
| WO1999007374A1 (en) * | 1997-08-04 | 1999-02-18 | Merck Sharp & Dohme Limited | Use of nk-1 receptor antagonists for treating aggressive behaviour |
| WO1999007376A1 (en) * | 1997-08-04 | 1999-02-18 | Merck Sharp & Dohme Limited | Use of nk-1 receptor antagonists for treating mania |
| WO1999007375A1 (en) * | 1997-08-04 | 1999-02-18 | Merck Sharp & Dohme Limited | Use of nk-1 receptor antagonists for treating aggressive behaviour disorders |
| WO1999007373A1 (en) * | 1997-08-04 | 1999-02-18 | Merck Sharp & Dohme Limited | Use of nk-1 receptor antagonists for treating mania |
| US5977104A (en) * | 1996-12-02 | 1999-11-02 | Merck Sharp & Dohme Ltd. | Use of NK-1 receptor antagonists for treating bipolar disorders |
| WO1999064010A1 (en) * | 1998-06-11 | 1999-12-16 | Merck Sharp & Dohme Limited | Use of a nk-1 receptor antagonist for treating psychiatric disorders |
| WO1999064007A1 (en) * | 1998-06-11 | 1999-12-16 | Merck Sharp & Dohme Limited | Use of a nk-1 receptor antagonist for treating bipolar disorders |
| WO1999064006A1 (en) * | 1998-06-11 | 1999-12-16 | Merck Sharp & Dohme Limited | Use of a nk-1 receptor antagonist for treating psychiatric disorders |
| WO1999064009A1 (en) * | 1998-06-11 | 1999-12-16 | Merck Sharp & Dohme Limited | Use of an nk-1 receptor antagonist for treating cognitive disorders |
| WO1999064008A1 (en) * | 1998-06-11 | 1999-12-16 | Merck Sharp & Dohme Limited | Use of a nk-1 receptor antagonist for treating sexual dysfunctions |
| WO1999065900A1 (en) * | 1998-06-16 | 1999-12-23 | Merck Sharp & Dohme Limited | Chemical synthesis of morpholine derivatives |
| WO2000007598A1 (en) * | 1998-08-04 | 2000-02-17 | Merck Sharp & Dohme Limited | Use of a nk-1 receptor antagonist for treating or preventing abnormal bone resorption |
| WO2000026215A1 (en) * | 1998-10-30 | 2000-05-11 | Merck & Co., Inc. | Process for the preparation of 2-(r)-(1- (r)-(3,5-bis (trifluoromethyl) phenyl)ethoxy) -4-((5-dimethylaminomethyl) -1,2,3-triazol -4-yl)methyl) -3-(s)-(4- fluorophenyl) morpholine |
| WO2000026214A1 (en) * | 1998-10-30 | 2000-05-11 | Merck & Co., Inc. | Linear process for the preparation of a morpholine compound |
| US6087348A (en) * | 1997-12-01 | 2000-07-11 | Merck Sharp & Dohme Ltd. | Use of NK-1 receptor antagonists for treating stress disorders |
| US6100256A (en) * | 1996-12-02 | 2000-08-08 | Merck Sharp & Dohme Ltd. | Use of NK-1 receptors antagonists for treating schizophrenic disorders |
| US6114315A (en) * | 1996-12-02 | 2000-09-05 | Merck Sharp & Dohme Ltd. | Use of NK-1 receptor antagonists for treating major depressive disorders with anxiety |
| US6117855A (en) * | 1996-10-07 | 2000-09-12 | Merck Sharp & Dohme Ltd. | Use of a NK-1 receptor antagonist and an antidepressant and/or an anti-anxiety agent |
| US6156749A (en) * | 1997-12-01 | 2000-12-05 | Merck Sharp & Dohme Limited | Use of NK-1 receptor antagonists for treating movement disorders |
| US6225316B1 (en) | 1999-02-24 | 2001-05-01 | Hoffmann-La Roche Inc. | 3-phenyl-pyridine-derivatives |
| US6271230B1 (en) | 1997-12-01 | 2001-08-07 | Merck Sharp & Dohme Limited | Use of NK-1 receptor antagonists for treating cognitive disorders |
| US6274588B1 (en) | 1999-05-31 | 2001-08-14 | Hoffmann-La Roche Inc. | 4-phenyl-pyrimidine derivatives |
| US6291465B1 (en) | 1999-03-09 | 2001-09-18 | Hoffmann-La Roche Inc. | Biphenyl derivatives |
| US6297375B1 (en) | 1999-02-24 | 2001-10-02 | Hoffmann-La Roche Inc. | 4-phenyl-pyridine derivatives |
| US6407111B1 (en) | 1999-02-24 | 2002-06-18 | Hoffmann-La Roche Inc. | Phenyl substituted pyridine and benzene derivatives |
| US6452001B2 (en) | 2000-05-25 | 2002-09-17 | Hoffmann-La Roche Inc. | Diazapane derivatives useful as antagonists of neurokinin 1 receptor and methods for their formation |
| US6482829B2 (en) | 2000-06-08 | 2002-11-19 | Hoffmann-La Roche Inc. | Substituted heterocyclic siprodecane compound active as an antagonist of neurokinin 1 receptor |
| US6576762B2 (en) | 2000-07-24 | 2003-06-10 | Hoffmann-La Roche Inc. | Heteroaromatic substituted amides with antagonistic activity to neurokinin 1 receptors |
| US6593472B2 (en) | 2000-07-14 | 2003-07-15 | Hoffman-La Roche Inc. | NK-1 receptor active amine oxide prodrugs |
| US6613765B1 (en) | 1996-12-02 | 2003-09-02 | Merck Sharp & Dohme Limited | Use of NK-1 receptor antagonists for treating major depressive disorders |
| US6638981B2 (en) | 2001-08-17 | 2003-10-28 | Epicept Corporation | Topical compositions and methods for treating pain |
| US6642226B2 (en) | 2001-02-06 | 2003-11-04 | Hoffman-La Roche Inc. | Substituted phenyl-piperidine methanone compounds |
| US6756380B1 (en) | 1999-05-31 | 2004-06-29 | Hoffmann-La Roche Inc. | 5-phenyl-pyrimidine derivatives |
| US6770637B2 (en) | 2000-08-08 | 2004-08-03 | Hoffmann-La Roche Inc. | Substituted 4-phenyl-pyridine compounds with activity as antagonists of neurokinin 1 receptors |
| US6787539B2 (en) | 2000-11-22 | 2004-09-07 | Hoffmann-La Roche Inc. | 2,4,5,-trisubstituted pyrimidine derivatives |
| US6849624B2 (en) | 2001-07-31 | 2005-02-01 | Hoffmann-La Roche Inc. | Aromatic and heteroaromatic substituted amides |
| WO2005077366A1 (en) | 2004-02-11 | 2005-08-25 | Munoz Saez Miguel | Use of non-peptidic nk1 receptor antagonists for the production of apoptosis in tumour cells |
| BG64622B1 (en) * | 1999-11-29 | 2005-09-30 | F. Hoffmann-La Roche Ag | 2-(3,5-bis-trifluoromethyl-phenyl)-n-methyl-n-(6-morpholine-4-yl-4-o-tolyl-pyridine-3-yl)-isobutyramide, method for its preparation and use |
| US7084152B2 (en) | 2000-07-11 | 2006-08-01 | Amr Technology, Inc. | 4-phenyl substituted tetrahydroisoquinolines therapeutic use thereof |
| WO2006123182A2 (en) | 2005-05-17 | 2006-11-23 | Merck Sharp & Dohme Limited | Cyclohexyl sulphones for treatment of cancer |
| US7160881B2 (en) | 2003-01-31 | 2007-01-09 | Hoffmann-La Roche Inc. | Crystalline modifications of 2-(3,5-bis-trifluoromethyl-phenyl)-N-[6-(1,1-dioxo-1 λ6-thiomorpholin-4-yl)-4-(4-fluoro-2-methyl-phenyl)-pyridin-3-yl]-N-methyl-isobutyramide |
| US7163949B1 (en) | 1999-11-03 | 2007-01-16 | Amr Technology, Inc. | 4-phenyl substituted tetrahydroisoquinolines and use thereof |
| WO2007011820A2 (en) | 2005-07-15 | 2007-01-25 | Amr Technology, Inc. | Aryl-and heteroaryl-substituted tetrahydrobenzazepines and use thereof to block reuptake of norepinephrine, dopamine, and serotonin |
| US7176205B2 (en) | 2005-02-22 | 2007-02-13 | Hoffmann-La Roche Inc. | Bi-pyridinyl derivatives as NK-1 antagonists |
| WO2007041052A2 (en) | 2005-09-29 | 2007-04-12 | Merck & Co., Inc. | Acylated spiropiperidine derivatives as melanocortin-4 receptor modulators |
| US7211579B2 (en) | 2005-03-23 | 2007-05-01 | Hoffmann-La Roche Inc. | NK-1 receptor antagonists |
| WO2007093827A1 (en) | 2006-02-15 | 2007-08-23 | Istituto Di Ricerche Di Biologia Molecolare P. Angeletti Spa | Thiophene and thiazole substituted trifluoroethanone derivatives as histone deacetylase (hdac) inhibitors |
| US7265116B2 (en) | 1999-11-03 | 2007-09-04 | Arm Technology, Inc. | Aryl and heteroaryl substituted tetrahydroisoquinolines and use thereof |
| US7288658B2 (en) | 2003-07-15 | 2007-10-30 | Hoffmann-La Roche Inc. | Process for preparation of pyridine derivatives |
| WO2008090117A1 (en) | 2007-01-24 | 2008-07-31 | Glaxo Group Limited | Pharmaceutical compositions comprising 3, 5-diamin0-6- (2, 3-dichl0phenyl) -l, 2, 4-triazine or r (-) -2, 4-diamino-5- (2, 3-dichlorophenyl) -6-fluoromethyl pyrimidine and an nk1 |
| WO2008120653A1 (en) | 2007-04-02 | 2008-10-09 | Banyu Pharmaceutical Co., Ltd. | Indoledione derivative |
| WO2009002495A1 (en) | 2007-06-27 | 2008-12-31 | Merck & Co., Inc. | 4-carboxybenzylamino derivatives as histone deacetylase inhibitors |
| US7541357B2 (en) | 2004-07-15 | 2009-06-02 | Amr Technology, Inc. | Aryl- and heteroaryl-substituted tetrahydroisoquinolines and use thereof to block reuptake of norepinephrine, dopamine, and serotonin |
| WO2009104080A2 (en) | 2008-02-20 | 2009-08-27 | Targia Pharmaceuticals | Cns pharmaceutical compositions and methods of use |
| WO2009111354A2 (en) | 2008-03-03 | 2009-09-11 | Tiger Pharmatech | Tyrosine kinase inhibitors |
| WO2010114780A1 (en) | 2009-04-01 | 2010-10-07 | Merck Sharp & Dohme Corp. | Inhibitors of akt activity |
| WO2010132442A1 (en) | 2009-05-12 | 2010-11-18 | Albany Molecular Reserch, Inc. | 7-([1,2,4,]triazolo[1,5,-a]pyridin-6-yl)-4-(3,4-dichlorophenyl)-1,2,3,4- tetrahydroisoquinoline and use thereof |
| WO2010132487A1 (en) | 2009-05-12 | 2010-11-18 | Bristol-Myers Squibb Company | CRYSTALLINE FORMS OF (S)-7-([1,2,4]TRIAZOLO[1,5-a]PYRIDIN-6-YL)-4-(3,4-DICHLOROHPHENYL)-1,2,3,4-TETRAHYDROISOQUINOLINE AND USE THEREOF |
| EP2266980A1 (en) * | 2005-10-05 | 2010-12-29 | Ranbaxy Laboratories Limited | Polymorphic form of aprepitant intermediate |
| US7893062B2 (en) | 2007-04-20 | 2011-02-22 | Hoffmann-La Roche Inc. | Pyrrolidine derivatives as dual NK1/NK3 receptor antagonists |
| WO2011046771A1 (en) | 2009-10-14 | 2011-04-21 | Schering Corporation | SUBSTITUTED PIPERIDINES THAT INCREASE p53 ACTIVITY AND THE USES THEREOF |
| US7939533B2 (en) | 2003-07-03 | 2011-05-10 | Hoffmann-La Roche Inc. | Dual NK1/NK3 receptor antagonists |
| EP2336120A1 (en) | 2007-01-10 | 2011-06-22 | Istituto di ricerche di Biologia Molecolare P. Angeletti S.R.L. | Combinations containing amide substituted indazoles as poly(ADP-ribose)polymerase (PARP) inhibitors |
| US8012998B2 (en) | 2007-08-07 | 2011-09-06 | Hoffmann—La Roche Inc. | Pyrrolidine aryl-ether as NK-3 receptor antagonists |
| WO2011131571A1 (en) | 2010-04-20 | 2011-10-27 | F. Hoffmann-La Roche Ag | Pyrrazolopyridine derivatives |
| WO2011163330A1 (en) | 2010-06-24 | 2011-12-29 | Merck Sharp & Dohme Corp. | Novel heterocyclic compounds as erk inhibitors |
| WO2012018754A2 (en) | 2010-08-02 | 2012-02-09 | Merck Sharp & Dohme Corp. | RNA INTERFERENCE MEDIATED INHIBITION OF CATENIN (CADHERIN-ASSOCIATED PROTEIN), BETA 1 (CTNNB1) GENE EXPRESSION USING SHORT INTERFERING NUCLEIC ACID (siNA) |
| WO2012020162A1 (en) | 2010-08-12 | 2012-02-16 | Salinas Martin Manuel Vicente | Use of antibodies against nk1, nk2 and/or nk3 receptors in cancer treatment |
| WO2012027236A1 (en) | 2010-08-23 | 2012-03-01 | Schering Corporation | NOVEL PYRAZOLO[1,5-a]PYRIMIDINE DERIVATIVES AS mTOR INHIBITORS |
| WO2012030685A2 (en) | 2010-09-01 | 2012-03-08 | Schering Corporation | Indazole derivatives useful as erk inhibitors |
| WO2012036997A1 (en) | 2010-09-16 | 2012-03-22 | Schering Corporation | Fused pyrazole derivatives as novel erk inhibitors |
| WO2012087772A1 (en) | 2010-12-21 | 2012-06-28 | Schering Corporation | Indazole derivatives useful as erk inhibitors |
| WO2012145471A1 (en) | 2011-04-21 | 2012-10-26 | Merck Sharp & Dohme Corp. | Insulin-like growth factor-1 receptor inhibitors |
| WO2013063214A1 (en) | 2011-10-27 | 2013-05-02 | Merck Sharp & Dohme Corp. | Novel compounds that are erk inhibitors |
| WO2013087964A1 (en) | 2011-12-13 | 2013-06-20 | Servicio Andaluz De Salud | Use of agents that alter the peritumoral environment for the treatment of cancer |
| WO2013165816A2 (en) | 2012-05-02 | 2013-11-07 | Merck Sharp & Dohme Corp. | SHORT INTERFERING NUCLEIC ACID (siNA) COMPOSITIONS |
| EP2698157A1 (en) | 2006-09-22 | 2014-02-19 | Merck Sharp & Dohme Corp. | Method of treatment using fatty acid synthesis inhibitors |
| WO2014052563A2 (en) | 2012-09-28 | 2014-04-03 | Merck Sharp & Dohme Corp. | Novel compounds that are erk inhibitors |
| WO2014085216A1 (en) | 2012-11-28 | 2014-06-05 | Merck Sharp & Dohme Corp. | Compositions and methods for treating cancer |
| WO2014100065A1 (en) | 2012-12-20 | 2014-06-26 | Merck Sharp & Dohme Corp. | Substituted imidazopyridines as hdm2 inhibitors |
| WO2014120748A1 (en) | 2013-01-30 | 2014-08-07 | Merck Sharp & Dohme Corp. | 2,6,7,8 substituted purines as hdm2 inhibitors |
| WO2014122353A1 (en) | 2013-02-11 | 2014-08-14 | Servicio Andaluz De Salud | Method for the prediction or prognosis of the response of a human subject suffering from cancer to treatment with an antagonist of the nk1 receptor |
| US8852634B2 (en) | 2005-09-23 | 2014-10-07 | Hoffmann-La Roche Inc. | Dosage formulation |
| WO2015034925A1 (en) | 2013-09-03 | 2015-03-12 | Moderna Therapeutics, Inc. | Circular polynucleotides |
| US9034899B2 (en) | 2009-05-12 | 2015-05-19 | Albany Molecular Research, Inc. | Aryl, heteroaryl, and heterocycle substituted tetrahydroisoquinolines and use thereof |
| WO2015097322A1 (en) | 2013-12-27 | 2015-07-02 | Servicio Andaluz De Salud | Use of non-peptide nk1 antagonists in a predetermined dose for the treatment of cancer |
| US9156812B2 (en) | 2008-06-04 | 2015-10-13 | Bristol-Myers Squibb Company | Crystalline form of 6-[(4S)-2-methyl-4-(2-naphthyl)-1,2,3,4-tetrahydroisoquinolin-7-yl]pyridazin-3-amine |
| WO2016046638A2 (en) | 2014-09-26 | 2016-03-31 | Helsinn Healthcare Sa | Crystalline forms of an nk-1 antagonist |
| WO2015101596A3 (en) * | 2013-12-30 | 2016-04-07 | Oncoprevent Gmbh | Neurokinin-1 receptor antagonists for use in a method of prevention of cancer |
| WO2017060338A1 (en) | 2015-10-06 | 2017-04-13 | Helsinn Healthcare Sa | Crystalline forms of fosnetupitant |
| WO2017211663A1 (en) | 2016-06-06 | 2017-12-14 | Helsinn Healthcare Sa | Physiologically balanced injectable formulations of fosnetupitant |
| WO2018071283A1 (en) | 2016-10-12 | 2018-04-19 | Merck Sharp & Dohme Corp. | Kdm5 inhibitors |
| EP3327125A1 (en) | 2010-10-29 | 2018-05-30 | Sirna Therapeutics, Inc. | Rna interference mediated inhibition of gene expression using short interfering nucleic acids (sina) |
| WO2019094311A1 (en) | 2017-11-08 | 2019-05-16 | Merck Sharp & Dohme Corp. | Prmt5 inhibitors |
| WO2019094312A1 (en) | 2017-11-08 | 2019-05-16 | Merck Sharp & Dohme Corp. | Prmt5 inhibitors |
| WO2020033288A1 (en) | 2018-08-07 | 2020-02-13 | Merck Sharp & Dohme Corp. | Prmt5 inhibitors |
| WO2020033284A1 (en) | 2018-08-07 | 2020-02-13 | Merck Sharp & Dohme Corp. | Prmt5 inhibitors |
| WO2020033282A1 (en) | 2018-08-07 | 2020-02-13 | Merck Sharp & Dohme Corp. | Prmt5 inhibitors |
| US11096950B2 (en) | 2006-11-01 | 2021-08-24 | Barbara Brooke Jennings | Compounds, methods, and treatments for abnormal signaling pathways for prenatal and postnatal development |
| EP4079856A1 (en) | 2010-08-17 | 2022-10-26 | Sirna Therapeutics, Inc. | Rna interference mediated inhibition of hepatitis b virus (hbv) gene expression using short interfering nucleic acid (sina) |
Families Citing this family (15)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| ES2157337T3 (en) * | 1994-08-15 | 2001-08-16 | Merck Sharp & Dohme | DERIVATIVES OF MORFOLINA AND ITS USE AS THERAPEUTIC AGENTS. |
| US5891875A (en) * | 1996-05-20 | 1999-04-06 | Eli Lilly And Company | Morpholinyl tachykinin receptor antagonists |
| US6274604B1 (en) | 1996-07-17 | 2001-08-14 | Merck & Co., Inc. | Alteration of circadian rhythmicity with a tachykinin antagonist |
| GB9700555D0 (en) * | 1997-01-13 | 1997-03-05 | Merck Sharp & Dohme | Therapeutic agents |
| US5972938A (en) * | 1997-12-01 | 1999-10-26 | Merck & Co., Inc. | Method for treating or preventing psychoimmunological disorders |
| GB9810092D0 (en) * | 1998-05-11 | 1998-07-08 | Merck Sharp & Dohme | Therapeutic agents |
| JP2003513095A (en) | 1999-10-29 | 2003-04-08 | メルク エンド カムパニー インコーポレーテッド | Polymorphs of tachykinin receptor antagonists |
| AR030284A1 (en) | 2000-06-08 | 2003-08-20 | Merck & Co Inc | PROCESS FOR THE SYNTHESIS OF (2R, 2-ALFA-R, 3A) -2- [1- (3,5-BIS (TRIFLUOROMETIL) PHENYL) ETOXI] -3- (4-FLUOROPHENIL) -1,4-OXAZINE; SUCH COMPOUND AND ITS POLYMORPHIC FORMS |
| WO2001096319A1 (en) * | 2000-06-09 | 2001-12-20 | Merck & Co., Inc. | Trans-glycosidation process for the synthesis of (2r, 2-alpha-r, 3a)-2-[1-(3,5-bis(trifluoromethyl)phenyl)ethoxy]-3-(4-fluorophenyl)-1,4-oxazine |
| GB0020721D0 (en) * | 2000-08-22 | 2000-10-11 | Merck Sharp & Dohme | Therapeutic agents |
| DE60217493T2 (en) * | 2001-07-16 | 2007-11-15 | Ortho-Mcneil Pharmaceutical, Inc. | USE OF CARBAMATE COMPOUNDS FOR THE PREVENTION OR TREATMENT OF NEUROPATHIC PAIN AND MIGRANT HEADACHE |
| AR039625A1 (en) * | 2002-04-18 | 2005-03-02 | Merck & Co Inc | PROCESS FOR THE PREPARATION OF 5 - ((2 (R) - (1 (R) - (3,5-BIS (TRIFLUORMETIL) PHENYL) ETOXI-3 (S) - (4-FLUORFENIL) -4-MORFOLINIL) METHYL ) -1,2-DIHIDRO-3H-1,2,4-TRIAZOL-3-ONA |
| RU2228752C1 (en) * | 2003-03-21 | 2004-05-20 | Государственный научный центр - Институт биофизики | Pharmaceutical composition containing ondansetron and benzamide |
| US8481565B2 (en) | 2004-12-27 | 2013-07-09 | Eisai R&D Management Co., Ltd. | Method for stabilizing anti-dementia drug |
| RU2392277C2 (en) * | 2005-12-28 | 2010-06-20 | Асино Фарма Аг | Method of nebivolol racemate obtainment |
Citations (2)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| EP0528495A1 (en) * | 1991-08-20 | 1993-02-24 | Merck Sharp & Dohme Ltd. | Azacyclic compounds, processes for their preparation and pharmaceutical compositions containing them |
| EP0577394A1 (en) * | 1992-06-29 | 1994-01-05 | Merck & Co. Inc. | Morpholine and thiomorpholine tachykinin receptor antagonists |
Family Cites Families (3)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| FR2534915B1 (en) * | 1982-10-26 | 1985-10-25 | Lafon Labor | NOVEL 2- (PHENOXYMETHYL) -MORPHOLINE DERIVATIVES, USE IN THERAPEUTICS AND METHOD OF PREPARATION |
| IL111960A (en) * | 1993-12-17 | 1999-12-22 | Merck & Co Inc | Morpholines and thiomorpholines their preparation and pharmaceutical compositions containing them |
| IL112778A0 (en) * | 1994-03-04 | 1995-05-26 | Merck & Co Inc | Substituted heterocycles, their preparation and pharmaceutical compositions containing them |
-
1994
- 1994-12-23 DE DE69434063T patent/DE69434063D1/en not_active Expired - Lifetime
- 1994-12-23 PL PL94315182A patent/PL181214B1/en not_active IP Right Cessation
- 1994-12-23 AT AT95904621T patent/ATE207917T1/en not_active IP Right Cessation
- 1994-12-23 CN CN94194722A patent/CN1067683C/en not_active Expired - Fee Related
- 1994-12-23 EP EP04076888A patent/EP1462450B1/en not_active Expired - Lifetime
- 1994-12-23 HU HU9601801A patent/HU221139B1/en not_active IP Right Cessation
- 1994-12-23 US US08/663,201 patent/US5612337A/en not_active Expired - Fee Related
- 1994-12-23 RU RU96116158/04A patent/RU2131426C1/en not_active IP Right Cessation
- 1994-12-23 CZ CZ19961898A patent/CZ288176B6/en not_active IP Right Cessation
- 1994-12-23 SK SK839-96A patent/SK283070B6/en unknown
- 1994-12-23 NZ NZ277839A patent/NZ277839A/en unknown
- 1994-12-23 DE DE69434991T patent/DE69434991T2/en not_active Expired - Fee Related
- 1994-12-23 SG SG1996000578A patent/SG52217A1/en unknown
- 1994-12-23 DE DE69428913T patent/DE69428913T2/en not_active Expired - Fee Related
- 1994-12-23 DK DK95904621T patent/DK0737192T3/en active
- 1994-12-23 BR BR9408442A patent/BR9408442A/en not_active Application Discontinuation
- 1994-12-23 CA CA002178219A patent/CA2178219C/en not_active Expired - Fee Related
- 1994-12-23 KR KR1019960703426A patent/KR100286412B1/en not_active Expired - Fee Related
- 1994-12-23 WO PCT/GB1994/002819 patent/WO1995018124A1/en not_active Ceased
- 1994-12-23 JP JP7517856A patent/JP3071467B2/en not_active Expired - Lifetime
- 1994-12-23 EP EP95904621A patent/EP0737192B1/en not_active Expired - Lifetime
- 1994-12-23 EP EP01200308A patent/EP1099702B1/en not_active Expired - Lifetime
- 1994-12-23 AT AT01200308T patent/ATE278687T1/en not_active IP Right Cessation
- 1994-12-23 AT AT04076888T patent/ATE364606T1/en not_active IP Right Cessation
- 1994-12-23 IL IL11213494A patent/IL112134A/en not_active IP Right Cessation
- 1994-12-23 ES ES95904621T patent/ES2164758T3/en not_active Expired - Lifetime
-
1995
- 1995-03-20 TW TW084102683A patent/TW449588B/en active
-
1996
- 1996-06-06 BG BG100644A patent/BG62616B1/en unknown
- 1996-06-19 LV LVP-96-197A patent/LV11687B/en unknown
- 1996-06-28 NO NO962749A patent/NO307090B1/en unknown
-
2000
- 2000-01-18 JP JP2000009217A patent/JP2000219629A/en active Pending
Patent Citations (2)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| EP0528495A1 (en) * | 1991-08-20 | 1993-02-24 | Merck Sharp & Dohme Ltd. | Azacyclic compounds, processes for their preparation and pharmaceutical compositions containing them |
| EP0577394A1 (en) * | 1992-06-29 | 1994-01-05 | Merck & Co. Inc. | Morpholine and thiomorpholine tachykinin receptor antagonists |
Cited By (171)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| US5719147A (en) * | 1992-06-29 | 1998-02-17 | Merck & Co., Inc. | Morpholine and thiomorpholine tachykinin receptor antagonists |
| WO1995030674A1 (en) * | 1994-05-05 | 1995-11-16 | Merck Sharp & Dohme Limited | Morpholine derivatives and their use as antagonists of tachikinins |
| US5747491A (en) * | 1994-05-05 | 1998-05-05 | Merck Sharp & Dohme Ltd. | Morpholine derivatives and their use as antagonists of tachikinins |
| US5719149A (en) * | 1994-09-02 | 1998-02-17 | Merck Sharp & Dohme Ltd. | Morpholine derivatives and their use as therapeutic agents |
| WO1996007649A1 (en) * | 1994-09-02 | 1996-03-14 | Merck Sharp & Dohme Limited | Morpholine derivatives and their use as therapeutic agents |
| US5880123A (en) * | 1994-12-23 | 1999-03-09 | Merck Sharp & Dohme Limited | Hemolysis prevention by non-ionic surfactants |
| WO1996019970A1 (en) * | 1994-12-23 | 1996-07-04 | Merck Sharp & Dohme Limited | Hemolysis prevention by surfactants and emulsions |
| WO1996029328A1 (en) * | 1995-03-18 | 1996-09-26 | Merck Sharp & Dohme Limited | Morpholine derivatives, compositions containing them and their use as therapeutic agents |
| US5877316A (en) * | 1995-03-18 | 1999-03-02 | Merck Sharp & Dohme Ltd. | Morpholine derivatives, compositions containing them and their use as therapeutic agents |
| WO1997002824A1 (en) * | 1995-07-08 | 1997-01-30 | Merck Sharp & Dohme Limited | Use of a tachykinin antagonist and a muscarinic antagonist and/or an antihistamine to treat motion sickness |
| US6140324A (en) * | 1995-07-08 | 2000-10-31 | Merck Sharp & Dohme Ltd. | Use of a tachykinin antagonist and a muscarinic antagonist and/or an antihistamine to treat motion sickness |
| WO1997018206A1 (en) * | 1995-11-14 | 1997-05-22 | Merck Sharp & Dohme Limited | Morpholine derivatives and their use as therapeutic agents |
| US5968934A (en) * | 1995-11-14 | 1999-10-19 | Merck Sharp & Dohme Ltd. | Morpholine derivatives and their use as therapeutic agents |
| US6117855A (en) * | 1996-10-07 | 2000-09-12 | Merck Sharp & Dohme Ltd. | Use of a NK-1 receptor antagonist and an antidepressant and/or an anti-anxiety agent |
| US6319953B1 (en) | 1996-10-07 | 2001-11-20 | Merck Sharp & Dohme Ltd. | Treatment of depression and anxiety with fluoxetine and an NK-1 receptor antagonist |
| US6649614B2 (en) | 1996-10-07 | 2003-11-18 | Merck Sharp & Dohme Ltd. | Use of a NK-1 receptor antagonist and an antidepressant and/or an anti-anxiety agent for treatment of depression or anxiety |
| WO1998024446A1 (en) * | 1996-12-02 | 1998-06-11 | Merck Sharp & Dohme Limited | Use of nk-1 receptor antagonists for treating movement disorders |
| WO1998024447A1 (en) * | 1996-12-02 | 1998-06-11 | Merck Sharp & Dohme Limited | Use of nk-1 receptor antagonists for treating cognitive disorders |
| US6096729A (en) * | 1996-12-02 | 2000-08-01 | Merck Sharp & Dohme Ltd. | Use of NK-1 receptor antagonists for treating amnestic disorders |
| US6100256A (en) * | 1996-12-02 | 2000-08-08 | Merck Sharp & Dohme Ltd. | Use of NK-1 receptors antagonists for treating schizophrenic disorders |
| WO1998024438A1 (en) * | 1996-12-02 | 1998-06-11 | Merck Sharp & Dohme Limited | Use of nk-1 receptor antagonists for treating major depressive disorders |
| WO1998024441A1 (en) * | 1996-12-02 | 1998-06-11 | Merck Sharp & Dohme Limited | Use of nk-1 receptor antagonists for treating major depressive disorders with anxiety |
| US6613765B1 (en) | 1996-12-02 | 2003-09-02 | Merck Sharp & Dohme Limited | Use of NK-1 receptor antagonists for treating major depressive disorders |
| WO1998024440A1 (en) * | 1996-12-02 | 1998-06-11 | Merck Sharp & Dohme Limited | Use of nk-1 receptor antagonists for treating stress disorders |
| WO1998024443A1 (en) * | 1996-12-02 | 1998-06-11 | Merck Sharp & Dohme Limited | Use of nk-1 receptor antagonists for treating bipolar disorders |
| WO1998024445A1 (en) * | 1996-12-02 | 1998-06-11 | Merck Sharp & Dohme Limited | Use of nk-1 receptor antagonists for treating schizophrenic disorders |
| US5919781A (en) * | 1996-12-02 | 1999-07-06 | Merck Sharp & Dohme Ltd. | Use of NK-1 receptor antogonists for treating substance use disorders |
| US5925627A (en) * | 1996-12-02 | 1999-07-20 | Merck Sharp & Dohme Ltd. | Use of NK-1 receptor antagonists for treating movement disorders |
| US5929054A (en) * | 1996-12-02 | 1999-07-27 | Merck Sharp | Use of NK-1 receptor antagonists for treating sexual dysfunctions |
| US5952315A (en) * | 1996-12-02 | 1999-09-14 | Merck Sharp & Dohme Ltd. | Use of NK-1 receptor antagonists for treating bipolar disorders |
| WO1998024439A1 (en) * | 1996-12-02 | 1998-06-11 | Merck Sharp & Dohme Limited | Use of nk-1 receptor antagonists for treating severe anxiety disorders |
| US5977104A (en) * | 1996-12-02 | 1999-11-02 | Merck Sharp & Dohme Ltd. | Use of NK-1 receptor antagonists for treating bipolar disorders |
| WO1998024444A1 (en) * | 1996-12-02 | 1998-06-11 | Merck Sharp & Dohme Limited | Use of nk-1 receptor antagonists for treating substance use disorders |
| WO1998024442A1 (en) * | 1996-12-02 | 1998-06-11 | Merck Sharp & Dohme Limited | Use of nk-1 receptor antagonists for treating sexual dysfunction |
| US6114315A (en) * | 1996-12-02 | 2000-09-05 | Merck Sharp & Dohme Ltd. | Use of NK-1 receptor antagonists for treating major depressive disorders with anxiety |
| WO1998047513A1 (en) * | 1997-04-24 | 1998-10-29 | Merck Sharp & Dohme Limited | Use of nk-1 receptor antagonists for treating eating disorders |
| WO1999007375A1 (en) * | 1997-08-04 | 1999-02-18 | Merck Sharp & Dohme Limited | Use of nk-1 receptor antagonists for treating aggressive behaviour disorders |
| WO1999007374A1 (en) * | 1997-08-04 | 1999-02-18 | Merck Sharp & Dohme Limited | Use of nk-1 receptor antagonists for treating aggressive behaviour |
| WO1999007376A1 (en) * | 1997-08-04 | 1999-02-18 | Merck Sharp & Dohme Limited | Use of nk-1 receptor antagonists for treating mania |
| WO1999007373A1 (en) * | 1997-08-04 | 1999-02-18 | Merck Sharp & Dohme Limited | Use of nk-1 receptor antagonists for treating mania |
| AU738047B2 (en) * | 1997-08-04 | 2001-09-06 | Merck Sharp & Dohme Limited | Use of NK-1 receptor antagonists for treating mania |
| US6271230B1 (en) | 1997-12-01 | 2001-08-07 | Merck Sharp & Dohme Limited | Use of NK-1 receptor antagonists for treating cognitive disorders |
| US6087348A (en) * | 1997-12-01 | 2000-07-11 | Merck Sharp & Dohme Ltd. | Use of NK-1 receptor antagonists for treating stress disorders |
| US6156749A (en) * | 1997-12-01 | 2000-12-05 | Merck Sharp & Dohme Limited | Use of NK-1 receptor antagonists for treating movement disorders |
| WO1999064008A1 (en) * | 1998-06-11 | 1999-12-16 | Merck Sharp & Dohme Limited | Use of a nk-1 receptor antagonist for treating sexual dysfunctions |
| WO1999064009A1 (en) * | 1998-06-11 | 1999-12-16 | Merck Sharp & Dohme Limited | Use of an nk-1 receptor antagonist for treating cognitive disorders |
| WO1999064006A1 (en) * | 1998-06-11 | 1999-12-16 | Merck Sharp & Dohme Limited | Use of a nk-1 receptor antagonist for treating psychiatric disorders |
| WO1999064007A1 (en) * | 1998-06-11 | 1999-12-16 | Merck Sharp & Dohme Limited | Use of a nk-1 receptor antagonist for treating bipolar disorders |
| WO1999064010A1 (en) * | 1998-06-11 | 1999-12-16 | Merck Sharp & Dohme Limited | Use of a nk-1 receptor antagonist for treating psychiatric disorders |
| US6407255B2 (en) | 1998-06-16 | 2002-06-18 | Merck Sharp & Dohme Ltd. | Chemical synthesis of 1,2,4-triazolinone derivative |
| WO1999065900A1 (en) * | 1998-06-16 | 1999-12-23 | Merck Sharp & Dohme Limited | Chemical synthesis of morpholine derivatives |
| HRP20000878B1 (en) * | 1998-06-16 | 2009-09-30 | Merck Sharp & Dohme Limited | CHEMICAL SYNTHESIS OF MORFOLIN DERIVATIVES |
| CZ299257B6 (en) * | 1998-06-16 | 2008-05-28 | Merck Sharp & Dohme Limited | Process for preparing morpholine derivatives |
| US6297376B1 (en) | 1998-06-16 | 2001-10-02 | Merck Sharp & Dohme Ltd. | Chemical synthesis of morpholine derivatives |
| AU746721B2 (en) * | 1998-06-16 | 2002-05-02 | Merck Sharp & Dohme Limited | Chemical synthesis of morpholine derivatives |
| WO2000007598A1 (en) * | 1998-08-04 | 2000-02-17 | Merck Sharp & Dohme Limited | Use of a nk-1 receptor antagonist for treating or preventing abnormal bone resorption |
| AU763615B2 (en) * | 1998-08-04 | 2003-07-31 | Merck Sharp & Dohme Limited | Use of a NK-1 receptor antagonist for treating or preventing abnormal bone resorption |
| WO2000026215A1 (en) * | 1998-10-30 | 2000-05-11 | Merck & Co., Inc. | Process for the preparation of 2-(r)-(1- (r)-(3,5-bis (trifluoromethyl) phenyl)ethoxy) -4-((5-dimethylaminomethyl) -1,2,3-triazol -4-yl)methyl) -3-(s)-(4- fluorophenyl) morpholine |
| WO2000026214A1 (en) * | 1998-10-30 | 2000-05-11 | Merck & Co., Inc. | Linear process for the preparation of a morpholine compound |
| US6225316B1 (en) | 1999-02-24 | 2001-05-01 | Hoffmann-La Roche Inc. | 3-phenyl-pyridine-derivatives |
| US6479483B2 (en) | 1999-02-24 | 2002-11-12 | Hoffmann-La Roche Inc. | 4-phenyl-pyridine derivatives |
| US6407111B1 (en) | 1999-02-24 | 2002-06-18 | Hoffmann-La Roche Inc. | Phenyl substituted pyridine and benzene derivatives |
| US6297375B1 (en) | 1999-02-24 | 2001-10-02 | Hoffmann-La Roche Inc. | 4-phenyl-pyridine derivatives |
| US6596773B2 (en) | 1999-02-24 | 2003-07-22 | Hoffmann-La Roche Inc. | Phenyl substituted pyridine and benzene derivatives |
| US6291465B1 (en) | 1999-03-09 | 2001-09-18 | Hoffmann-La Roche Inc. | Biphenyl derivatives |
| US6552088B2 (en) | 1999-03-09 | 2003-04-22 | Hoffmann-La Roche Inc. | Biphenyl derivatives |
| US6756380B1 (en) | 1999-05-31 | 2004-06-29 | Hoffmann-La Roche Inc. | 5-phenyl-pyrimidine derivatives |
| US6274588B1 (en) | 1999-05-31 | 2001-08-14 | Hoffmann-La Roche Inc. | 4-phenyl-pyrimidine derivatives |
| US7265116B2 (en) | 1999-11-03 | 2007-09-04 | Arm Technology, Inc. | Aryl and heteroaryl substituted tetrahydroisoquinolines and use thereof |
| US7612090B2 (en) | 1999-11-03 | 2009-11-03 | Albany Molecular Research, Inc. | Aryl and heteroaryl substituted tetrahydroisoquinolines and use thereof |
| US7163949B1 (en) | 1999-11-03 | 2007-01-16 | Amr Technology, Inc. | 4-phenyl substituted tetrahydroisoquinolines and use thereof |
| CZ299286B6 (en) * | 1999-11-29 | 2008-06-04 | F. Hoffmann-La Roche Ag | 2-(3,5-bis-Trifluoromethyl-phenyl)-N-methyl-N-(6-morpholin-4-yl-4-o-tolyl-pyridin-3-yl)-isobutyramide, process of its preparation and medicament containing this compound |
| BG64622B1 (en) * | 1999-11-29 | 2005-09-30 | F. Hoffmann-La Roche Ag | 2-(3,5-bis-trifluoromethyl-phenyl)-n-methyl-n-(6-morpholine-4-yl-4-o-tolyl-pyridine-3-yl)-isobutyramide, method for its preparation and use |
| US6452001B2 (en) | 2000-05-25 | 2002-09-17 | Hoffmann-La Roche Inc. | Diazapane derivatives useful as antagonists of neurokinin 1 receptor and methods for their formation |
| US6482829B2 (en) | 2000-06-08 | 2002-11-19 | Hoffmann-La Roche Inc. | Substituted heterocyclic siprodecane compound active as an antagonist of neurokinin 1 receptor |
| US7419985B2 (en) | 2000-07-11 | 2008-09-02 | Amr Technology, Inc. | 4-Phenyl substituted tetrahydroisoquinolines and therapeutic use thereof |
| US7309789B2 (en) | 2000-07-11 | 2007-12-18 | Amr Technology, Inc. | 4-phenyl substituted tetrahydroisoquinolines and therapeutic use thereof |
| US7084152B2 (en) | 2000-07-11 | 2006-08-01 | Amr Technology, Inc. | 4-phenyl substituted tetrahydroisoquinolines therapeutic use thereof |
| US6593472B2 (en) | 2000-07-14 | 2003-07-15 | Hoffman-La Roche Inc. | NK-1 receptor active amine oxide prodrugs |
| US6897226B2 (en) | 2000-07-14 | 2005-05-24 | Hoffmann-La Roche Inc. | NK-1 receptor active amine oxide prodrugs |
| US6806370B2 (en) | 2000-07-14 | 2004-10-19 | Hoffmann-La Roche Inc. | NK-1 receptor active amine oxide prodrugs |
| US6747026B2 (en) | 2000-07-14 | 2004-06-08 | Hoffman-La Roche Inc. | NK-1 receptor active amine oxide prodrugs |
| US6624176B2 (en) | 2000-07-24 | 2003-09-23 | Hoffmann-La Roche Inc. | Heteroaromatic substituted amides with antagonistic activity to neurokinin 1 receptors |
| US6576762B2 (en) | 2000-07-24 | 2003-06-10 | Hoffmann-La Roche Inc. | Heteroaromatic substituted amides with antagonistic activity to neurokinin 1 receptors |
| US6770637B2 (en) | 2000-08-08 | 2004-08-03 | Hoffmann-La Roche Inc. | Substituted 4-phenyl-pyridine compounds with activity as antagonists of neurokinin 1 receptors |
| US6787539B2 (en) | 2000-11-22 | 2004-09-07 | Hoffmann-La Roche Inc. | 2,4,5,-trisubstituted pyrimidine derivatives |
| US6642226B2 (en) | 2001-02-06 | 2003-11-04 | Hoffman-La Roche Inc. | Substituted phenyl-piperidine methanone compounds |
| US6849624B2 (en) | 2001-07-31 | 2005-02-01 | Hoffmann-La Roche Inc. | Aromatic and heteroaromatic substituted amides |
| US6638981B2 (en) | 2001-08-17 | 2003-10-28 | Epicept Corporation | Topical compositions and methods for treating pain |
| US7160881B2 (en) | 2003-01-31 | 2007-01-09 | Hoffmann-La Roche Inc. | Crystalline modifications of 2-(3,5-bis-trifluoromethyl-phenyl)-N-[6-(1,1-dioxo-1 λ6-thiomorpholin-4-yl)-4-(4-fluoro-2-methyl-phenyl)-pyridin-3-yl]-N-methyl-isobutyramide |
| RU2330022C2 (en) * | 2003-01-31 | 2008-07-27 | Ф.Хоффманн-Ля Рош Аг | NEW CRYSTALLINE MODIFICATION 2-(3, 5-BIS-TRIFLUOROMETHYLPHENYL)-H-[6-(1, 1-DIOXO-1 λ6 -THIOMORPHOLINE-4-YL)-4-(4-FLUORO-2-MEHTYLPHENYL)PIRIDINE-3-YL]-K-METHYLISOBUTYR AMIDE |
| US7939533B2 (en) | 2003-07-03 | 2011-05-10 | Hoffmann-La Roche Inc. | Dual NK1/NK3 receptor antagonists |
| US8404708B2 (en) | 2003-07-03 | 2013-03-26 | Hoffmann-La Roche Inc. | Dual NK1/NK3 receptor antagonists |
| US7288658B2 (en) | 2003-07-15 | 2007-10-30 | Hoffmann-La Roche Inc. | Process for preparation of pyridine derivatives |
| WO2005077366A1 (en) | 2004-02-11 | 2005-08-25 | Munoz Saez Miguel | Use of non-peptidic nk1 receptor antagonists for the production of apoptosis in tumour cells |
| EP2842605A2 (en) | 2004-02-11 | 2015-03-04 | Miguel Munoz Sáez | Use of non-peptide nk1 receptor antagonists for the production of apoptosis in tumor cells |
| US9499531B2 (en) | 2004-07-15 | 2016-11-22 | Albany Molecular Research, Inc. | Aryl- and heteroaryl-substituted tetrahydroisoquinolines and use thereof to block reuptake of norepinephrine, dopamine, and serotonin |
| US7541357B2 (en) | 2004-07-15 | 2009-06-02 | Amr Technology, Inc. | Aryl- and heteroaryl-substituted tetrahydroisoquinolines and use thereof to block reuptake of norepinephrine, dopamine, and serotonin |
| US9085531B2 (en) | 2004-07-15 | 2015-07-21 | Albany Molecular Research, Inc. | Aryl- and heteroaryl-substituted tetrahydroisoquinolines and use thereof to block reuptake of norepinephrine, dopamine, and serotonin |
| US7176205B2 (en) | 2005-02-22 | 2007-02-13 | Hoffmann-La Roche Inc. | Bi-pyridinyl derivatives as NK-1 antagonists |
| US7211579B2 (en) | 2005-03-23 | 2007-05-01 | Hoffmann-La Roche Inc. | NK-1 receptor antagonists |
| WO2006123182A2 (en) | 2005-05-17 | 2006-11-23 | Merck Sharp & Dohme Limited | Cyclohexyl sulphones for treatment of cancer |
| WO2007011820A2 (en) | 2005-07-15 | 2007-01-25 | Amr Technology, Inc. | Aryl-and heteroaryl-substituted tetrahydrobenzazepines and use thereof to block reuptake of norepinephrine, dopamine, and serotonin |
| US9403776B2 (en) | 2005-07-15 | 2016-08-02 | Albany Molecular Research, Inc. | Aryl- and heteroaryl-substituted tetrahydrobenzazepines and use thereof to block reuptake of norepinephrine, dopamine, and serotonin |
| US8852634B2 (en) | 2005-09-23 | 2014-10-07 | Hoffmann-La Roche Inc. | Dosage formulation |
| WO2007041052A2 (en) | 2005-09-29 | 2007-04-12 | Merck & Co., Inc. | Acylated spiropiperidine derivatives as melanocortin-4 receptor modulators |
| EP2266980A1 (en) * | 2005-10-05 | 2010-12-29 | Ranbaxy Laboratories Limited | Polymorphic form of aprepitant intermediate |
| US8080656B2 (en) | 2005-10-05 | 2011-12-20 | Ranbaxy Laboratories Limited | Process for the preparation of aprepitant |
| WO2007093827A1 (en) | 2006-02-15 | 2007-08-23 | Istituto Di Ricerche Di Biologia Molecolare P. Angeletti Spa | Thiophene and thiazole substituted trifluoroethanone derivatives as histone deacetylase (hdac) inhibitors |
| EP2946778A1 (en) | 2006-09-22 | 2015-11-25 | Merck Sharp & Dohme Corp. | Method of treatment using fatty acid synthesis inhibitors |
| EP2698157A1 (en) | 2006-09-22 | 2014-02-19 | Merck Sharp & Dohme Corp. | Method of treatment using fatty acid synthesis inhibitors |
| US11096950B2 (en) | 2006-11-01 | 2021-08-24 | Barbara Brooke Jennings | Compounds, methods, and treatments for abnormal signaling pathways for prenatal and postnatal development |
| EP2336120A1 (en) | 2007-01-10 | 2011-06-22 | Istituto di ricerche di Biologia Molecolare P. Angeletti S.R.L. | Combinations containing amide substituted indazoles as poly(ADP-ribose)polymerase (PARP) inhibitors |
| EP2805945A1 (en) | 2007-01-10 | 2014-11-26 | MSD Italia S.r.l. | Amide substituted indazoles as poly(ADP-ribose)polymerase (PARP) inhibitors |
| WO2008090117A1 (en) | 2007-01-24 | 2008-07-31 | Glaxo Group Limited | Pharmaceutical compositions comprising 3, 5-diamin0-6- (2, 3-dichl0phenyl) -l, 2, 4-triazine or r (-) -2, 4-diamino-5- (2, 3-dichlorophenyl) -6-fluoromethyl pyrimidine and an nk1 |
| WO2008120653A1 (en) | 2007-04-02 | 2008-10-09 | Banyu Pharmaceutical Co., Ltd. | Indoledione derivative |
| US7893062B2 (en) | 2007-04-20 | 2011-02-22 | Hoffmann-La Roche Inc. | Pyrrolidine derivatives as dual NK1/NK3 receptor antagonists |
| EP3103791A1 (en) | 2007-06-27 | 2016-12-14 | Merck Sharp & Dohme Corp. | 4-carboxybenzylamino derivatives as histone deacetylase inhibitors |
| WO2009002495A1 (en) | 2007-06-27 | 2008-12-31 | Merck & Co., Inc. | 4-carboxybenzylamino derivatives as histone deacetylase inhibitors |
| US8012998B2 (en) | 2007-08-07 | 2011-09-06 | Hoffmann—La Roche Inc. | Pyrrolidine aryl-ether as NK-3 receptor antagonists |
| WO2009104080A2 (en) | 2008-02-20 | 2009-08-27 | Targia Pharmaceuticals | Cns pharmaceutical compositions and methods of use |
| WO2009111354A2 (en) | 2008-03-03 | 2009-09-11 | Tiger Pharmatech | Tyrosine kinase inhibitors |
| US9156812B2 (en) | 2008-06-04 | 2015-10-13 | Bristol-Myers Squibb Company | Crystalline form of 6-[(4S)-2-methyl-4-(2-naphthyl)-1,2,3,4-tetrahydroisoquinolin-7-yl]pyridazin-3-amine |
| US9498476B2 (en) | 2008-06-04 | 2016-11-22 | Albany Molecular Research, Inc. | Crystalline form of 6-[(4S)-2-methyl-4-(2-naphthyl)-1,2,3,4-tetrahydroisoquinolin-7-yl]pyridazin-3-amine |
| WO2010114780A1 (en) | 2009-04-01 | 2010-10-07 | Merck Sharp & Dohme Corp. | Inhibitors of akt activity |
| US9604960B2 (en) | 2009-05-12 | 2017-03-28 | Albany Molecular Research, Inc. | Aryl, heteroaryl, and heterocycle substituted tetrahydroisoquinolines and use thereof |
| WO2010132442A1 (en) | 2009-05-12 | 2010-11-18 | Albany Molecular Reserch, Inc. | 7-([1,2,4,]triazolo[1,5,-a]pyridin-6-yl)-4-(3,4-dichlorophenyl)-1,2,3,4- tetrahydroisoquinoline and use thereof |
| US9173879B2 (en) | 2009-05-12 | 2015-11-03 | Bristol-Myers Squibb Company | Crystalline forms of (S)-7-([1,2,4]triazolo[1,5-a ]pyridin-6-yl)-4-(3,4-dichlorophenyl)-1,2,3,4-tetrahydroisoquinoline and use thereof |
| WO2010132487A1 (en) | 2009-05-12 | 2010-11-18 | Bristol-Myers Squibb Company | CRYSTALLINE FORMS OF (S)-7-([1,2,4]TRIAZOLO[1,5-a]PYRIDIN-6-YL)-4-(3,4-DICHLOROHPHENYL)-1,2,3,4-TETRAHYDROISOQUINOLINE AND USE THEREOF |
| US9034899B2 (en) | 2009-05-12 | 2015-05-19 | Albany Molecular Research, Inc. | Aryl, heteroaryl, and heterocycle substituted tetrahydroisoquinolines and use thereof |
| WO2011046771A1 (en) | 2009-10-14 | 2011-04-21 | Schering Corporation | SUBSTITUTED PIPERIDINES THAT INCREASE p53 ACTIVITY AND THE USES THEREOF |
| WO2011131571A1 (en) | 2010-04-20 | 2011-10-27 | F. Hoffmann-La Roche Ag | Pyrrazolopyridine derivatives |
| US8487102B2 (en) | 2010-04-20 | 2013-07-16 | Hoffmann-La Roche Inc. | Pyrrazolopyridine compounds as dual NK1/NK3 receptor antagonists |
| WO2011163330A1 (en) | 2010-06-24 | 2011-12-29 | Merck Sharp & Dohme Corp. | Novel heterocyclic compounds as erk inhibitors |
| EP3330377A1 (en) | 2010-08-02 | 2018-06-06 | Sirna Therapeutics, Inc. | Rna interference mediated inhibition of catenin (cadherin-associated protein), beta 1 (ctnnb1) gene expression using short interfering nucleic acid (sina) |
| WO2012018754A2 (en) | 2010-08-02 | 2012-02-09 | Merck Sharp & Dohme Corp. | RNA INTERFERENCE MEDIATED INHIBITION OF CATENIN (CADHERIN-ASSOCIATED PROTEIN), BETA 1 (CTNNB1) GENE EXPRESSION USING SHORT INTERFERING NUCLEIC ACID (siNA) |
| WO2012020162A1 (en) | 2010-08-12 | 2012-02-16 | Salinas Martin Manuel Vicente | Use of antibodies against nk1, nk2 and/or nk3 receptors in cancer treatment |
| EP4079856A1 (en) | 2010-08-17 | 2022-10-26 | Sirna Therapeutics, Inc. | Rna interference mediated inhibition of hepatitis b virus (hbv) gene expression using short interfering nucleic acid (sina) |
| WO2012027236A1 (en) | 2010-08-23 | 2012-03-01 | Schering Corporation | NOVEL PYRAZOLO[1,5-a]PYRIMIDINE DERIVATIVES AS mTOR INHIBITORS |
| WO2012030685A2 (en) | 2010-09-01 | 2012-03-08 | Schering Corporation | Indazole derivatives useful as erk inhibitors |
| WO2012036997A1 (en) | 2010-09-16 | 2012-03-22 | Schering Corporation | Fused pyrazole derivatives as novel erk inhibitors |
| EP3327125A1 (en) | 2010-10-29 | 2018-05-30 | Sirna Therapeutics, Inc. | Rna interference mediated inhibition of gene expression using short interfering nucleic acids (sina) |
| EP3766975A1 (en) | 2010-10-29 | 2021-01-20 | Sirna Therapeutics, Inc. | Rna interference mediated inhibition of gene expression using short interfering nucleic acid (sina) |
| WO2012087772A1 (en) | 2010-12-21 | 2012-06-28 | Schering Corporation | Indazole derivatives useful as erk inhibitors |
| WO2012145471A1 (en) | 2011-04-21 | 2012-10-26 | Merck Sharp & Dohme Corp. | Insulin-like growth factor-1 receptor inhibitors |
| WO2013063214A1 (en) | 2011-10-27 | 2013-05-02 | Merck Sharp & Dohme Corp. | Novel compounds that are erk inhibitors |
| WO2013087964A1 (en) | 2011-12-13 | 2013-06-20 | Servicio Andaluz De Salud | Use of agents that alter the peritumoral environment for the treatment of cancer |
| EP3919620A1 (en) | 2012-05-02 | 2021-12-08 | Sirna Therapeutics, Inc. | Short interfering nucleic acid (sina) compositions |
| WO2013165816A2 (en) | 2012-05-02 | 2013-11-07 | Merck Sharp & Dohme Corp. | SHORT INTERFERING NUCLEIC ACID (siNA) COMPOSITIONS |
| WO2014052563A2 (en) | 2012-09-28 | 2014-04-03 | Merck Sharp & Dohme Corp. | Novel compounds that are erk inhibitors |
| WO2014085216A1 (en) | 2012-11-28 | 2014-06-05 | Merck Sharp & Dohme Corp. | Compositions and methods for treating cancer |
| WO2014100065A1 (en) | 2012-12-20 | 2014-06-26 | Merck Sharp & Dohme Corp. | Substituted imidazopyridines as hdm2 inhibitors |
| WO2014120748A1 (en) | 2013-01-30 | 2014-08-07 | Merck Sharp & Dohme Corp. | 2,6,7,8 substituted purines as hdm2 inhibitors |
| WO2014122353A1 (en) | 2013-02-11 | 2014-08-14 | Servicio Andaluz De Salud | Method for the prediction or prognosis of the response of a human subject suffering from cancer to treatment with an antagonist of the nk1 receptor |
| WO2015034925A1 (en) | 2013-09-03 | 2015-03-12 | Moderna Therapeutics, Inc. | Circular polynucleotides |
| WO2015097322A1 (en) | 2013-12-27 | 2015-07-02 | Servicio Andaluz De Salud | Use of non-peptide nk1 antagonists in a predetermined dose for the treatment of cancer |
| WO2015101596A3 (en) * | 2013-12-30 | 2016-04-07 | Oncoprevent Gmbh | Neurokinin-1 receptor antagonists for use in a method of prevention of cancer |
| WO2016046638A2 (en) | 2014-09-26 | 2016-03-31 | Helsinn Healthcare Sa | Crystalline forms of an nk-1 antagonist |
| US10005803B2 (en) | 2015-10-06 | 2018-06-26 | Helsinn Healthcare Sa | Crystalline forms of fosnetupitant |
| WO2017060338A1 (en) | 2015-10-06 | 2017-04-13 | Helsinn Healthcare Sa | Crystalline forms of fosnetupitant |
| US12208109B2 (en) | 2016-06-06 | 2025-01-28 | Helsinn Healthcare Sa | Physiologically balanced injectable formulations of fosnetupitant |
| US11529362B2 (en) | 2016-06-06 | 2022-12-20 | Helsinn Healthcare Sa | Physiologically balanced injectable formulations of fosnetupitant |
| WO2017211663A1 (en) | 2016-06-06 | 2017-12-14 | Helsinn Healthcare Sa | Physiologically balanced injectable formulations of fosnetupitant |
| EP3626231A1 (en) | 2016-06-06 | 2020-03-25 | Helsinn Healthcare SA | Physiologically balanced injectable formulations of fosnetupitant |
| US10624911B2 (en) | 2016-06-06 | 2020-04-21 | Helsinn Healthcare Sa | Physiologically balanced injectable formulations of fosnetupitant |
| WO2018071283A1 (en) | 2016-10-12 | 2018-04-19 | Merck Sharp & Dohme Corp. | Kdm5 inhibitors |
| WO2019094312A1 (en) | 2017-11-08 | 2019-05-16 | Merck Sharp & Dohme Corp. | Prmt5 inhibitors |
| WO2019094311A1 (en) | 2017-11-08 | 2019-05-16 | Merck Sharp & Dohme Corp. | Prmt5 inhibitors |
| WO2020033282A1 (en) | 2018-08-07 | 2020-02-13 | Merck Sharp & Dohme Corp. | Prmt5 inhibitors |
| WO2020033284A1 (en) | 2018-08-07 | 2020-02-13 | Merck Sharp & Dohme Corp. | Prmt5 inhibitors |
| WO2020033288A1 (en) | 2018-08-07 | 2020-02-13 | Merck Sharp & Dohme Corp. | Prmt5 inhibitors |
Also Published As
Similar Documents
| Publication | Publication Date | Title |
|---|---|---|
| EP0737192A1 (en) | Substituted morpholine derivatives and their use as therapeutic agents | |
| EP0777659B1 (en) | Morpholine derivatives and their use as therapeutic agents | |
| US5512570A (en) | Treatment of emesis with morpholine tachykinin receptor antagonists | |
| EP0734381B1 (en) | Morpholine tachykinin receptor antagonists | |
| US5610159A (en) | N-oxides of morpholine derivatives and their use as therapeutic agents | |
| US5719149A (en) | Morpholine derivatives and their use as therapeutic agents | |
| EP0577394A1 (en) | Morpholine and thiomorpholine tachykinin receptor antagonists | |
| US6638930B2 (en) | Morpholine and thiomorpholine tachykinin receptor antagonists | |
| AU685209C (en) | Substituted morpholine derivatives and their use as therapeutic agents | |
| HRP950222A2 (en) | Substituted morpholine derivatives and their use as therapeutic agents |
Legal Events
| Date | Code | Title | Description |
|---|---|---|---|
| WWE | Wipo information: entry into national phase |
Ref document number: 94194722.X Country of ref document: CN |
|
| AK | Designated states |
Kind code of ref document: A1 Designated state(s): AM AT AU BB BG BR BY CA CH CN CZ DE DK EE ES FI GB GE HU JP KE KG KP KR KZ LK LR LT LU LV MD MG MN MW NL NO NZ PL PT RO RU SD SE SI SK TJ TT UA US UZ VN |
|
| AL | Designated countries for regional patents |
Kind code of ref document: A1 Designated state(s): KE MW SD SZ AT BE CH DE DK ES FR GB GR IE IT LU MC NL PT SE BF BJ CF CG CI CM GA GN ML MR NE SN TD TG |
|
| DFPE | Request for preliminary examination filed prior to expiration of 19th month from priority date (pct application filed before 20040101) | ||
| 121 | Ep: the epo has been informed by wipo that ep was designated in this application | ||
| WWE | Wipo information: entry into national phase |
Ref document number: 277839 Country of ref document: NZ |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 1995904621 Country of ref document: EP |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 2178219 Country of ref document: CA |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 08663201 Country of ref document: US |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 83996 Country of ref document: SK |
|
| WWE | Wipo information: entry into national phase |
Ref document number: PV1996-1898 Country of ref document: CZ |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 96-01327 Country of ref document: RO |
|
| WWP | Wipo information: published in national office |
Ref document number: 1995904621 Country of ref document: EP |
|
| REG | Reference to national code |
Ref country code: DE Ref legal event code: 8642 |
|
| WWP | Wipo information: published in national office |
Ref document number: PV1996-1898 Country of ref document: CZ |
|
| WWG | Wipo information: grant in national office |
Ref document number: PV1996-1898 Country of ref document: CZ |
|
| WWG | Wipo information: grant in national office |
Ref document number: 1995904621 Country of ref document: EP |