WO1998052920A1 - Chemical process for the reduction of 1-substituted -3-hydroxymethyl-4- (4-fluorophenyl)tetrahydropyridines - Google Patents
Chemical process for the reduction of 1-substituted -3-hydroxymethyl-4- (4-fluorophenyl)tetrahydropyridines Download PDFInfo
- Publication number
- WO1998052920A1 WO1998052920A1 PCT/EP1998/002826 EP9802826W WO9852920A1 WO 1998052920 A1 WO1998052920 A1 WO 1998052920A1 EP 9802826 W EP9802826 W EP 9802826W WO 9852920 A1 WO9852920 A1 WO 9852920A1
- Authority
- WO
- WIPO (PCT)
- Prior art keywords
- hydride
- sodium
- process according
- aluminium hydride
- lithium
- Prior art date
- Legal status (The legal status is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the status listed.)
- Ceased
Links
- 0 *N(CC1)C[C@@](CO)[C@@]1c(cc1)ccc1F Chemical compound *N(CC1)C[C@@](CO)[C@@]1c(cc1)ccc1F 0.000 description 1
Classifications
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D405/00—Heterocyclic compounds containing both one or more hetero rings having oxygen atoms as the only ring hetero atoms, and one or more rings having nitrogen as the only ring hetero atom
- C07D405/02—Heterocyclic compounds containing both one or more hetero rings having oxygen atoms as the only ring hetero atoms, and one or more rings having nitrogen as the only ring hetero atom containing two hetero rings
- C07D405/12—Heterocyclic compounds containing both one or more hetero rings having oxygen atoms as the only ring hetero atoms, and one or more rings having nitrogen as the only ring hetero atom containing two hetero rings linked by a chain containing hetero atoms as chain links
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D211/00—Heterocyclic compounds containing hydrogenated pyridine rings, not condensed with other rings
- C07D211/04—Heterocyclic compounds containing hydrogenated pyridine rings, not condensed with other rings with only hydrogen or carbon atoms directly attached to the ring nitrogen atom
- C07D211/06—Heterocyclic compounds containing hydrogenated pyridine rings, not condensed with other rings with only hydrogen or carbon atoms directly attached to the ring nitrogen atom having no double bonds between ring members or between ring members and non-ring members
- C07D211/08—Heterocyclic compounds containing hydrogenated pyridine rings, not condensed with other rings with only hydrogen or carbon atoms directly attached to the ring nitrogen atom having no double bonds between ring members or between ring members and non-ring members with hydrocarbon or substituted hydrocarbon radicals directly attached to ring carbon atoms
- C07D211/18—Heterocyclic compounds containing hydrogenated pyridine rings, not condensed with other rings with only hydrogen or carbon atoms directly attached to the ring nitrogen atom having no double bonds between ring members or between ring members and non-ring members with hydrocarbon or substituted hydrocarbon radicals directly attached to ring carbon atoms with substituted hydrocarbon radicals attached to ring carbon atoms
- C07D211/20—Heterocyclic compounds containing hydrogenated pyridine rings, not condensed with other rings with only hydrogen or carbon atoms directly attached to the ring nitrogen atom having no double bonds between ring members or between ring members and non-ring members with hydrocarbon or substituted hydrocarbon radicals directly attached to ring carbon atoms with substituted hydrocarbon radicals attached to ring carbon atoms with hydrocarbon radicals, substituted by singly bound oxygen or sulphur atoms
- C07D211/22—Heterocyclic compounds containing hydrogenated pyridine rings, not condensed with other rings with only hydrogen or carbon atoms directly attached to the ring nitrogen atom having no double bonds between ring members or between ring members and non-ring members with hydrocarbon or substituted hydrocarbon radicals directly attached to ring carbon atoms with substituted hydrocarbon radicals attached to ring carbon atoms with hydrocarbon radicals, substituted by singly bound oxygen or sulphur atoms by oxygen atoms
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D211/00—Heterocyclic compounds containing hydrogenated pyridine rings, not condensed with other rings
- C07D211/04—Heterocyclic compounds containing hydrogenated pyridine rings, not condensed with other rings with only hydrogen or carbon atoms directly attached to the ring nitrogen atom
- C07D211/06—Heterocyclic compounds containing hydrogenated pyridine rings, not condensed with other rings with only hydrogen or carbon atoms directly attached to the ring nitrogen atom having no double bonds between ring members or between ring members and non-ring members
- C07D211/08—Heterocyclic compounds containing hydrogenated pyridine rings, not condensed with other rings with only hydrogen or carbon atoms directly attached to the ring nitrogen atom having no double bonds between ring members or between ring members and non-ring members with hydrocarbon or substituted hydrocarbon radicals directly attached to ring carbon atoms
- C07D211/18—Heterocyclic compounds containing hydrogenated pyridine rings, not condensed with other rings with only hydrogen or carbon atoms directly attached to the ring nitrogen atom having no double bonds between ring members or between ring members and non-ring members with hydrocarbon or substituted hydrocarbon radicals directly attached to ring carbon atoms with substituted hydrocarbon radicals attached to ring carbon atoms
- C07D211/34—Heterocyclic compounds containing hydrogenated pyridine rings, not condensed with other rings with only hydrogen or carbon atoms directly attached to the ring nitrogen atom having no double bonds between ring members or between ring members and non-ring members with hydrocarbon or substituted hydrocarbon radicals directly attached to ring carbon atoms with substituted hydrocarbon radicals attached to ring carbon atoms with hydrocarbon radicals, substituted by carbon atoms having three bonds to hetero atoms with at the most one bond to halogen, e.g. ester or nitrile radicals
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D211/00—Heterocyclic compounds containing hydrogenated pyridine rings, not condensed with other rings
- C07D211/04—Heterocyclic compounds containing hydrogenated pyridine rings, not condensed with other rings with only hydrogen or carbon atoms directly attached to the ring nitrogen atom
- C07D211/68—Heterocyclic compounds containing hydrogenated pyridine rings, not condensed with other rings with only hydrogen or carbon atoms directly attached to the ring nitrogen atom having one double bond between ring members or between a ring member and a non-ring member
- C07D211/70—Heterocyclic compounds containing hydrogenated pyridine rings, not condensed with other rings with only hydrogen or carbon atoms directly attached to the ring nitrogen atom having one double bond between ring members or between a ring member and a non-ring member with only hydrogen atoms, hydrocarbon or substituted hydrocarbon radicals, directly attached to ring carbon atoms
-
- Y—GENERAL TAGGING OF NEW TECHNOLOGICAL DEVELOPMENTS; GENERAL TAGGING OF CROSS-SECTIONAL TECHNOLOGIES SPANNING OVER SEVERAL SECTIONS OF THE IPC; TECHNICAL SUBJECTS COVERED BY FORMER USPC CROSS-REFERENCE ART COLLECTIONS [XRACs] AND DIGESTS
- Y02—TECHNOLOGIES OR APPLICATIONS FOR MITIGATION OR ADAPTATION AGAINST CLIMATE CHANGE
- Y02P—CLIMATE CHANGE MITIGATION TECHNOLOGIES IN THE PRODUCTION OR PROCESSING OF GOODS
- Y02P20/00—Technologies relating to chemical industry
- Y02P20/50—Improvements relating to the production of bulk chemicals
- Y02P20/55—Design of synthesis routes, e.g. reducing the use of auxiliary or protecting groups
Definitions
- This invention relates to a process for the preparation of (-)-trans-1 -benzyl-3- hydroxymethyl-4-(4-fluorophenyl)piperidine which is a useful intermediate in the preparation of paroxetine.
- This compound has been found to be especially useful in the treatment of depression and several processes have been described to prepare this important compound.
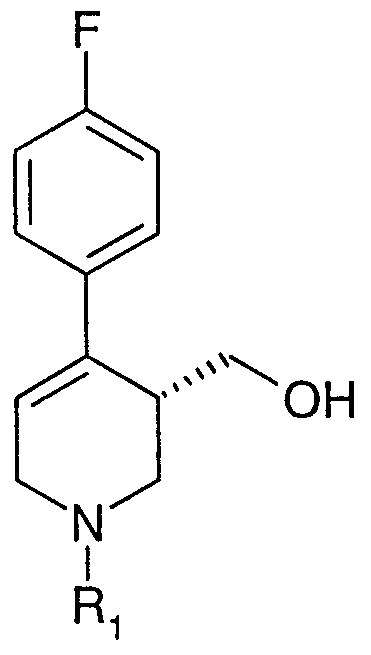
- Step C in claim 1 of this application describes the reduction of a compound of formula B
- a compound of formula C in which X is halogen, preferably F, and Ri is C 2 - 5 alkyl, phenyl C 1- alkyl or substituted phenyl C ⁇ - 5 alkyl.
- the reduction is carried out using a metal hydride which is preferably, according to claim 4, LiAIH 4 or NaAIH 4 .
- a compound of formula B in which X is F and R-i is ethyl, which was reduced using a mixture of sodium hydride and lithium aluminium hydride.
- the present invention provides a process for the preparation of a compound of formula I
- Ri is as defined above is reduced by a metal hydride in the presence of an inorganic salt in the presence of a diluent.
- the amine protecting group is one which is inert to reduction by a metal hydride.
- the amine protecting group is selected from a) allyl, b) benzhydryl, c) methoxymethyl, d) benzyloxymethyl, e) tetrahydropyranyl, f) an optionally substituted benzyl group, g) di(p-methoxyphenyl)methyl, h) triphenylmethyl, i) (p-methoxyphenyl)diphenylmethyl, j) diphenyl-4-pyridylmethyl, k) 2,4,6-trimethylbenzenesulphonyl, I) toluenesulphonyl, m) benzylsulphonyl, n) a C ⁇ alkyl group, o) a trifluoro C ⁇ -4 alkyl group, p) an alkynyl group or q) p-methoxybenzyl or
- the amine protecting group is a benzyl group which is optionally substituted on the phenyl ring by one or more of the following groups: a C-
- Ri represents benzyl.
- the metal hydride is sodium hydride, potassium hydride, magnesium hydride, calcium hydride, sodium borohydride, potassium borohydride, lithium borohydride, lithium aluminium hydride, sodium aluminium hydride, aluminium hydride, sodium bis-(2-methoxyethoxy)aluminium hydride, a lithium mono(C ⁇ -4 alkoxy)aluminium hydride, a lithium di(C ⁇ -4 alkoxy)aluminium hydride or sodium diethylaluminium hydride or mixtures thereof.
- the metal hydride is lithium aluminium hydride or sodium aluminium hydride. More preferably the metal hydride is lithium aluminium hydride.
- the amount of metal hydride used is in the range of 0.5 - 5 molar equivalents with respect to the amount of compound of formula II used.
- the amount of metal hydride used is in the range of 0.75 - 1.25 molar equivalents.
- the amount of metal hydride used is in the range of 0.90 - 1.10 molar equivalents.
- the inorganic salt is a salt of lithium, sodium, magnesium, calcium, zinc, lanthanum or iron, or mixtures thereof.
- the inorganic salt is a halide salt of lithium, sodium, calcium, zinc, magnesium, lanthanum or iron or mixtures thereof.
- the inorganic salt is selected from lithium chloride, sodium chloride, calcium chloride, zinc chloride, iron (II) chloride, iron (III) chloride, lanthanum chloride, magnesium chloride, magnesium fluoride, magnesium bromide or magnesium iodide or mixtures thereof.
- the inorganic salt is magnesium chloride, magnesium bromide or magnesium iodide.
- An especially preferred salt is magnesium chloride.
- the active reducing agent may be formed by a reaction between the metal hydride initially employed and the inorganic salt.
- the active species may be one or more of magnesium hydride, chloromagnesium aluminium hydride, magnesium aluminium hydride or lithium magnesium aluminium hydride or a complex of magnesium chloride and lithium aluminium hydride. It is to be understood that this process covers all such equivalents.
- the amount of inorganic salt used is in the range of 0.25 molar equivalents to 5 molar equivalents with respect to the amount of the compound of formula II used.
- the amount of inorganic salt used is in the range of 0.5 - 1.5. More preferably the amount of inorganic salt used is in the range of 0.75 -1.25 molar equivalents with respect to the amount of the compound of formula II used.
- the diluent is an organic liquid which is inert to the metal hydride employed and is preferably a solvent for the compound of formula II.
- the diluent is an ether or a hydrocarbon or a mixture thereof. More preferably the diluent is selected from tetrahydrofuran, toluene, dioxane, diethyl ether, diisopropyl ether, t-butylmethyl ether, diglyme, ethylene glycol dimethyl ether or mixtures thereof. Most preferably the diluent is tetrahydrofuran.
- the amount of the diluent is in the range of 1 part by weight to 100 parts by weight with respect to the compound of formula II employed.
- the amount of the diluent is in the range of 2 parts by weight to 50 parts by weight with respect to the compound of formula II employed. More preferably the amount of the diluent is in the range of 3 parts by weight to 10 parts by weight with respect to the compound of formula II employed.
- the process is carried out at a temperature in the range of -70°C to the boiling point of the diluent employed.
- the process is carried out at a temperature in the range 0-150°C. More preferably the process is carried out at a temperature in the range 0-100°C. Most preferably the process is carried out at a temperature in the range 50-70°C.
- the amount of desfluoro compound obtained is in the range of
- the amount of desfluoro compound obtained is in the range of 0.001 to 0.5%. More preferably the amount of desfluoro compound obtained is in the range of 0.001 to 0.2%.
- the process of the present invention is advantageous because it provides a pure precursor to paroxetine.
- Paroxetine may be obtained in a pure form from compounds of formula I by a) conversion of the hydroxy group into a leaving group, for example halo or tosyloxy, b) reaction with sesamol or a salt thereof, c) removal of the protecting group Ri and optionally d) salt formation, for example the hydrochloride salt as the anhydrous form or the hemihydrate.
- the invention is illustrated by the following Examples which are given by way of example only.
- the final product of each of these Examples was characterised by one or more of the following procedures: gas-liquid chromatography; high performance liquid chromatography; elemental analysis; nuclear magnetic resonance spectroscopy and infrared spectroscopy.
- the desfluoro compound is (-)-trans-1 -benzyl-3-hydroxymethyl-4- phenylpiperidine.
- Carrier gas (B) Flow: 4.5 mls/min. Initial temp: 40°C for 1 min.
- Examples 3-8 were carried out in a similar manner to Example 1 using the conditions described in Table 1.
- M equiv II represents molar equivalents with respect to the amount of the compound of formula II employed
- GLC gas liquid chromatography
- HPLC high performance liquid chromatography
- %(-F) % of desfluoro compound
Landscapes
- Chemical & Material Sciences (AREA)
- Organic Chemistry (AREA)
- Hydrogenated Pyridines (AREA)
- Nitrogen Condensed Heterocyclic Rings (AREA)
- Plural Heterocyclic Compounds (AREA)
- Pharmaceuticals Containing Other Organic And Inorganic Compounds (AREA)
- Low-Molecular Organic Synthesis Reactions Using Catalysts (AREA)
- Pyridine Compounds (AREA)
- Organic Low-Molecular-Weight Compounds And Preparation Thereof (AREA)
Abstract
Description


Claims

Priority Applications (18)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| DK98929307T DK0983237T3 (en) | 1997-05-17 | 1998-05-13 | Chemical Process for Reduction of 1-Substituted 3-Hydroxymethyl-4- (4-Fluorophenyl) Tetrahydro-Pyridines |
| EP98929307A EP0983237B1 (en) | 1997-05-17 | 1998-05-13 | Chemical process for the reduction of 1-substituted -3-hydroxymethyl-4- (4-fluorophenyl)tetrahydropyridines |
| PL98336796A PL189464B1 (en) | 1997-05-17 | 1998-05-13 | Chemical method of reducing 1-substituted 3-hydroxymethyl-4-(4-fluorophenyl)-tetrahydropyridines |
| JP54990898A JP4463883B2 (en) | 1997-05-17 | 1998-05-13 | Chemical reduction method for 1-substituted-3-hydroxymethyl-4- (4-fluorophenyl) tetrahydropyridine |
| BRPI9809809-8A BR9809809B1 (en) | 1997-05-17 | 1998-05-13 | process for the preparation of a (-) - trans-1-benzyl-3-hydroxymethyl-4- (4-fluorophenyl) piperidine compound; and, process for the preparation of paroxetine. |
| NZ501692A NZ501692A (en) | 1997-05-17 | 1998-05-13 | Chemical process for the reduction of 1-substituted -3-hydroxymethyl-4- (4-fluorophenyl)tetrahydropyridines, and use in preparing paroxetine |
| SK1495-99A SK149599A3 (en) | 1997-05-17 | 1998-05-13 | Chemical process for the reduction of 1-substituted-3- -hydroxymethyl-4-(4-fluorophenyl)tetrahydropyridines |
| SI9830095T SI0983237T1 (en) | 1997-05-17 | 1998-05-13 | Chemical process for the reduction of 1-substituted -3-hydroxymethyl-4- (4-fluorophenyl)tetrahydropyridines |
| KR10-1999-7010581A KR100515852B1 (en) | 1997-05-17 | 1998-05-13 | Chemical Process for the Reduction of 1-substituted-3-hydroxymethyl-4-(4-fluorophenyl)tetrahydropyridines |
| IL13236098A IL132360A (en) | 1997-05-17 | 1998-05-13 | Chemical process for the reduction of 1-substituted 3-hydroxymethyl-4-(4-fluorophenyl) tetrahydropyridines |
| UA99126877A UA61952C2 (en) | 1997-05-17 | 1998-05-13 | A chemical process for reduction of 1-substituted 3-hydroxymethyl-4-(4-fluorophenyl)tetrahydropyridines |
| DE69803178T DE69803178T2 (en) | 1997-05-17 | 1998-05-13 | METHOD FOR REDUCING 1-SUBSTITUTED-3-HYDROXYMETHYL-4- (4-FLUOROPHENYL) TETRAHYDROPYRIDINE |
| US09/423,749 US6326496B1 (en) | 1997-05-17 | 1998-05-13 | Process for preparing an intermediate in the production of paroxetine |
| CA002289013A CA2289013C (en) | 1997-05-17 | 1998-05-13 | Chemical process for the reduction of 1-substituted-3-hydroxymethyl-4-(4-fluorophenyl)tetrahydropyridines |
| AT98929307T ATE212013T1 (en) | 1997-05-17 | 1998-05-13 | METHOD FOR REDUCING 1-SUBSTITUTED-3-HYDROXYMETHYL-4-(4-FLUOROPHENYL) TETRAHYDROPYRIDINES |
| AU79121/98A AU748902B2 (en) | 1997-05-17 | 1998-05-13 | Chemical process for the reduction of 1-substituted -3-hydroxymethyl-4- (4-fluorophenyl)tetrahydropyridines |
| BG103834A BG63847B1 (en) | 1997-05-17 | 1999-10-26 | Method for reducing 1-substituted-3-hydroxymethyl-4-(4-fluorphenyl)tetrahydropyridines |
| NO19995619A NO313326B1 (en) | 1997-05-17 | 1999-11-16 | Chemical process for reduction of 1-substituted 2-hydroxymethyl-4- (4-fluorophenyl) tetrahydropyridines |
Applications Claiming Priority (2)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| GBGB9710004.4A GB9710004D0 (en) | 1997-05-17 | 1997-05-17 | Chemical process |
| GB9710004.4 | 1997-05-17 |
Publications (1)
| Publication Number | Publication Date |
|---|---|
| WO1998052920A1 true WO1998052920A1 (en) | 1998-11-26 |
Family
ID=10812456
Family Applications (1)
| Application Number | Title | Priority Date | Filing Date |
|---|---|---|---|
| PCT/EP1998/002826 Ceased WO1998052920A1 (en) | 1997-05-17 | 1998-05-13 | Chemical process for the reduction of 1-substituted -3-hydroxymethyl-4- (4-fluorophenyl)tetrahydropyridines |
Country Status (28)
| Country | Link |
|---|---|
| US (1) | US6326496B1 (en) |
| EP (1) | EP0983237B1 (en) |
| JP (1) | JP4463883B2 (en) |
| KR (1) | KR100515852B1 (en) |
| CN (1) | CN1121386C (en) |
| AR (1) | AR012688A1 (en) |
| AT (1) | ATE212013T1 (en) |
| AU (1) | AU748902B2 (en) |
| BG (1) | BG63847B1 (en) |
| BR (1) | BR9809809B1 (en) |
| CA (1) | CA2289013C (en) |
| CZ (1) | CZ292282B6 (en) |
| DE (1) | DE69803178T2 (en) |
| DK (1) | DK0983237T3 (en) |
| ES (1) | ES2171299T3 (en) |
| GB (1) | GB9710004D0 (en) |
| HU (1) | HUP0002709A3 (en) |
| ID (1) | ID24181A (en) |
| IL (1) | IL132360A (en) |
| NO (1) | NO313326B1 (en) |
| NZ (1) | NZ501692A (en) |
| PL (1) | PL189464B1 (en) |
| PT (1) | PT983237E (en) |
| RU (1) | RU2205178C2 (en) |
| SK (1) | SK149599A3 (en) |
| TR (1) | TR199902823T2 (en) |
| UA (1) | UA61952C2 (en) |
| WO (1) | WO1998052920A1 (en) |
Cited By (2)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| WO2001085688A1 (en) * | 2000-05-12 | 2001-11-15 | Synthon B.V. | TOSYLATE SALTS OF 4-(p-FLUOROPHENYL)-PIPERIDINE-3-CARBINOLS |
| US6949650B2 (en) | 2000-08-30 | 2005-09-27 | Aesica Pharmaceuticals Ltd. | Process for the racemization of 1-benzyl-4-(4-fluorophenyl)-3-hydroxymethyl-1,2-3,6-tetrahydropyridine to be used as intermediate in the synthesis of paroxetine |
Families Citing this family (6)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| US6777554B2 (en) | 2001-02-05 | 2004-08-17 | Teva Pharmaceutical Industries Ltd. | Preparation of N-methylparoxetine and related intermediate compounds |
| US6930186B2 (en) * | 2001-06-13 | 2005-08-16 | Teva Pharmacetical Industries Ltd. | Process for the preparation of paroxetine substantially free of alkoxy impurities |
| US20050266082A1 (en) * | 2004-05-26 | 2005-12-01 | Patel Satishkumar A | Preparation of stable paroxetine HC1 ER tablets using a melt granulation process |
| WO2007058998A2 (en) * | 2005-11-14 | 2007-05-24 | Auspex Pharmaceuticals, Inc. | Substituted phenylpiperidines with serotoninergic activity and enhanced therapeutic properties |
| US20080033050A1 (en) | 2006-08-04 | 2008-02-07 | Richards Patricia Allison Tewe | Method of treating thermoregulatory disfunction with paroxetine |
| US9138430B2 (en) * | 2007-12-27 | 2015-09-22 | Mylan Specialty L.P. | Formulation and method for the release of paroxetine in the large intestine |
Citations (2)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| US3849542A (en) * | 1966-05-09 | 1974-11-19 | Dow Chemical Co | Lithium magnesium aluminum hydride |
| WO1996036636A1 (en) * | 1995-05-17 | 1996-11-21 | Novo Nordisk A/S | Process for preparing 4-aryl-piperidine derivatives |
Family Cites Families (9)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| GB1422263A (en) | 1973-01-30 | 1976-01-21 | Ferrosan As | 4-phenyl-piperidine compounds |
| DE3680184D1 (en) * | 1985-08-10 | 1991-08-14 | Beecham Group Plc | METHOD FOR PRODUCING ARYLPIPERIDINE CARBINOL. |
| EP0299549A3 (en) * | 1987-07-09 | 1989-02-08 | Duphar International Research B.V | Tertiary 2,5-dialkyl-3-phenyl-piperidine derivatives having opiate-antagonistic activity |
| GB9100505D0 (en) * | 1991-01-10 | 1991-02-20 | Shell Int Research | Piperidine derivatives |
| US5258517A (en) * | 1992-08-06 | 1993-11-02 | Sepracor, Inc. | Method of preparing optically pure precursors of paroxetine |
| GB9305175D0 (en) * | 1993-03-13 | 1993-04-28 | Smithkline Beecham Plc | Novel process |
| EP1394160A1 (en) * | 1996-06-13 | 2004-03-03 | SUMIKA FINE CHEMICALS Co., Ltd. | Process for preparing crystalline paroxetin hydrochloride |
| HU221921B1 (en) | 1996-07-08 | 2003-02-28 | Richter Gedeon Vegyészeti Gyár Rt. | N-benzyl-piperidine or tetrahydro-pyridine derivatives and processes for producing them |
| GB9623359D0 (en) * | 1996-11-09 | 1997-01-08 | Smithkline Beecham Plc | Novel process |
-
1997
- 1997-05-17 GB GBGB9710004.4A patent/GB9710004D0/en active Pending
-
1998
- 1998-05-12 AR ARP980102206A patent/AR012688A1/en unknown
- 1998-05-13 UA UA99126877A patent/UA61952C2/en unknown
- 1998-05-13 PT PT98929307T patent/PT983237E/en unknown
- 1998-05-13 WO PCT/EP1998/002826 patent/WO1998052920A1/en not_active Ceased
- 1998-05-13 NZ NZ501692A patent/NZ501692A/en unknown
- 1998-05-13 US US09/423,749 patent/US6326496B1/en not_active Expired - Fee Related
- 1998-05-13 IL IL13236098A patent/IL132360A/en not_active IP Right Cessation
- 1998-05-13 ID IDW991271A patent/ID24181A/en unknown
- 1998-05-13 PL PL98336796A patent/PL189464B1/en not_active IP Right Cessation
- 1998-05-13 CZ CZ19994081A patent/CZ292282B6/en not_active IP Right Cessation
- 1998-05-13 KR KR10-1999-7010581A patent/KR100515852B1/en not_active Expired - Fee Related
- 1998-05-13 TR TR1999/02823T patent/TR199902823T2/en unknown
- 1998-05-13 CN CN98805113A patent/CN1121386C/en not_active Expired - Fee Related
- 1998-05-13 AU AU79121/98A patent/AU748902B2/en not_active Ceased
- 1998-05-13 DE DE69803178T patent/DE69803178T2/en not_active Expired - Lifetime
- 1998-05-13 CA CA002289013A patent/CA2289013C/en not_active Expired - Fee Related
- 1998-05-13 JP JP54990898A patent/JP4463883B2/en not_active Expired - Fee Related
- 1998-05-13 ES ES98929307T patent/ES2171299T3/en not_active Expired - Lifetime
- 1998-05-13 DK DK98929307T patent/DK0983237T3/en active
- 1998-05-13 AT AT98929307T patent/ATE212013T1/en not_active IP Right Cessation
- 1998-05-13 SK SK1495-99A patent/SK149599A3/en unknown
- 1998-05-13 BR BRPI9809809-8A patent/BR9809809B1/en not_active IP Right Cessation
- 1998-05-13 RU RU99127285/04A patent/RU2205178C2/en not_active IP Right Cessation
- 1998-05-13 EP EP98929307A patent/EP0983237B1/en not_active Expired - Lifetime
- 1998-05-13 HU HU0002709A patent/HUP0002709A3/en unknown
-
1999
- 1999-10-26 BG BG103834A patent/BG63847B1/en unknown
- 1999-11-16 NO NO19995619A patent/NO313326B1/en unknown
Patent Citations (2)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| US3849542A (en) * | 1966-05-09 | 1974-11-19 | Dow Chemical Co | Lithium magnesium aluminum hydride |
| WO1996036636A1 (en) * | 1995-05-17 | 1996-11-21 | Novo Nordisk A/S | Process for preparing 4-aryl-piperidine derivatives |
Non-Patent Citations (1)
| Title |
|---|
| WILLOCKS K ET AL: "THE SYNTHESIS OF 14C-3S,4R-4-(4-FLUOROPHENYL)-3- (3,4-METHYLENEDIOXYPHENOXYMETHYL)PIPERIDINE HYDROCHLORIDE (BRL 29060A), AND MECHANISTIC STUDIES USING CARBON-13 LABELLING", JOURNAL OF LABELLED COMPOUNDS AND RADIOPHARMACEUTICALS, vol. 33, no. 8, 1993, pages 783 - 794, XP000645839 * |
Cited By (2)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| WO2001085688A1 (en) * | 2000-05-12 | 2001-11-15 | Synthon B.V. | TOSYLATE SALTS OF 4-(p-FLUOROPHENYL)-PIPERIDINE-3-CARBINOLS |
| US6949650B2 (en) | 2000-08-30 | 2005-09-27 | Aesica Pharmaceuticals Ltd. | Process for the racemization of 1-benzyl-4-(4-fluorophenyl)-3-hydroxymethyl-1,2-3,6-tetrahydropyridine to be used as intermediate in the synthesis of paroxetine |
Also Published As
Similar Documents
| Publication | Publication Date | Title |
|---|---|---|
| EP0219934B1 (en) | Process for the preparation of aryl-piperidine esters | |
| EP0983237B1 (en) | Chemical process for the reduction of 1-substituted -3-hydroxymethyl-4- (4-fluorophenyl)tetrahydropyridines | |
| US20020198387A1 (en) | Process for producing paroxetine | |
| PL191995B1 (en) | Method of synthesis of 3-ethyl-5-methyl-2-(2-aminoethoxymethyl)- 4-(2-chlorophenyl)-6-methyl-1,4-dihydro-3,5-pyridine dicarbo-xylate and benzenesulfoacid salt, and their intermediates | |
| JPH0649683B2 (en) | Novel method for producing compound | |
| EP0437258B1 (en) | Method of manufacturing aromatic urethane | |
| EP0058069B1 (en) | Process and intermediates for preparing pirbuterol and analogs | |
| EP1124802A1 (en) | Process for producing 4-arylpiperidine-3-carbinols and related compounds | |
| EP0165682B1 (en) | The preparation of 1-(3,4-dimethoxyphenethyl amino)-3-(substituted phenoxy)-2-propanols | |
| US7323607B2 (en) | Process for preparation of (+)-p-mentha-2,8-diene-1-ol | |
| JP2006524247A5 (en) | ||
| EP0831081B1 (en) | Production of aminophenols | |
| EP3900724B1 (en) | Isoquinoline sulfonyl chloride acid addition salts and method of manufacture thereof | |
| JP2007001978A (en) | Method for producing ether | |
| EP0789573A1 (en) | Reductions using hydrides | |
| US20020095047A1 (en) | Process for preparing hydroxychomanones and CIS-aminochromanols | |
| JPH0625122A (en) | Aniline derivative and method for producing the same | |
| WO2002032870A1 (en) | Process of the preparation of 3-substituted-4-aryl piperidine compounds | |
| EP0650957A1 (en) | Aniline derivative and process for producing the same | |
| WO2002018337A1 (en) | Process for the racemisation of an intermediate useful in the preparation of paroxetine | |
| IE46154B1 (en) | Azetidinone derivatives |
Legal Events
| Date | Code | Title | Description |
|---|---|---|---|
| WWE | Wipo information: entry into national phase |
Ref document number: 132360 Country of ref document: IL Ref document number: 98805113.3 Country of ref document: CN |
|
| AK | Designated states |
Kind code of ref document: A1 Designated state(s): AL AU BG BR BY CA CN CZ GE HU ID IL JP KR KZ LT LV MX NO NZ PL RO RU SG SI SK TR UA US |
|
| AL | Designated countries for regional patents |
Kind code of ref document: A1 Designated state(s): AM AZ BY KG KZ MD RU TJ TM AT BE CH CY DE DK ES FI FR GB GR IE IT LU MC NL PT SE |
|
| DFPE | Request for preliminary examination filed prior to expiration of 19th month from priority date (pct application filed before 20040101) | ||
| 121 | Ep: the epo has been informed by wipo that ep was designated in this application | ||
| WWE | Wipo information: entry into national phase |
Ref document number: 1998929307 Country of ref document: EP |
|
| WWE | Wipo information: entry into national phase |
Ref document number: PA/a/1999/009667 Country of ref document: MX |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 149599 Country of ref document: SK |
|
| ENP | Entry into the national phase |
Ref document number: 2289013 Country of ref document: CA Ref document number: 2289013 Country of ref document: CA Kind code of ref document: A |
|
| ENP | Entry into the national phase |
Ref document number: 1998 549908 Country of ref document: JP Kind code of ref document: A |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 09423749 Country of ref document: US |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 79121/98 Country of ref document: AU Ref document number: 1019997010581 Country of ref document: KR |
|
| WWE | Wipo information: entry into national phase |
Ref document number: PV1999-4081 Country of ref document: CZ Ref document number: 1999/02823 Country of ref document: TR |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 501692 Country of ref document: NZ |
|
| WWP | Wipo information: published in national office |
Ref document number: 1998929307 Country of ref document: EP |
|
| WWP | Wipo information: published in national office |
Ref document number: PV1999-4081 Country of ref document: CZ |
|
| WWP | Wipo information: published in national office |
Ref document number: 1019997010581 Country of ref document: KR |
|
| WWG | Wipo information: grant in national office |
Ref document number: 1998929307 Country of ref document: EP |
|
| WWG | Wipo information: grant in national office |
Ref document number: 79121/98 Country of ref document: AU |
|
| WWG | Wipo information: grant in national office |
Ref document number: PV1999-4081 Country of ref document: CZ |
|
| WWG | Wipo information: grant in national office |
Ref document number: 1019997010581 Country of ref document: KR |