WO1999011269A1 - Pyrimidine derivatives - Google Patents

Pyrimidine derivatives Download PDF

Info

Publication number
WO1999011269A1
WO1999011269A1 PCT/US1998/018381 US9818381W WO9911269A1 WO 1999011269 A1 WO1999011269 A1 WO 1999011269A1 US 9818381 W US9818381 W US 9818381W WO 9911269 A1 WO9911269 A1 WO 9911269A1
Authority
WO
WIPO (PCT)
Prior art keywords
compound
formula
pharmaceutically acceptable
group
salt
Prior art date
Legal status (The legal status is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the status listed.)
Ceased
Application number
PCT/US1998/018381
Other languages
French (fr)
Inventor
Trent Lee Abraham
Boris Arnoldovich Czeskis
Minxia He
Lisa Ann Shipley
Current Assignee (The listed assignees may be inaccurate. Google has not performed a legal analysis and makes no representation or warranty as to the accuracy of the list.)
Eli Lilly and Co
Original Assignee
Eli Lilly and Co
Priority date (The priority date is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the date listed.)
Filing date
Publication date
Application filed by Eli Lilly and Co filed Critical Eli Lilly and Co
Priority to AU93022/98A priority Critical patent/AU9302298A/en
Priority to CA002269971A priority patent/CA2269971A1/en
Priority to EP98945876A priority patent/EP0954316A4/en
Publication of WO1999011269A1 publication Critical patent/WO1999011269A1/en
Anticipated expiration legal-status Critical
Ceased legal-status Critical Current

Links

Classifications

    • C—CHEMISTRY; METALLURGY
    • C07—ORGANIC CHEMISTRY
    • C07D—HETEROCYCLIC COMPOUNDS
    • C07D403/00—Heterocyclic compounds containing two or more hetero rings, having nitrogen atoms as the only ring hetero atoms, not provided for by group C07D401/00
    • C07D403/02—Heterocyclic compounds containing two or more hetero rings, having nitrogen atoms as the only ring hetero atoms, not provided for by group C07D401/00 containing two hetero rings
    • C07D403/12—Heterocyclic compounds containing two or more hetero rings, having nitrogen atoms as the only ring hetero atoms, not provided for by group C07D401/00 containing two hetero rings linked by a chain containing hetero atoms as chain links

Definitions

  • the present invention relates to novel pyrimidine derivatives, to processes for preparing them, to intermediates useful in preparing them, to pharmaceutical compositions containing them and to uses for them as pharmaceuticals .
  • United States patent number 4,323,570 discloses a group of aminopyrimidine derivatives having antihypertensive activity.
  • One of the compounds, 4-chloro-5- (imidazoline-2- ylamino) -6-methoxy-2-methylpyrimidine is commercially available in some countries under the generic name moxonidine for the treatment of hypertension.
  • This compound may interconvert with its tautomer, 4-chloro-5- (imidazolidin-2 -ylideneimino) - 6 -methoxy-2 -methylpyrimidine .
  • 4-chloro-5- (imidazolidin-2 -ylideneimino) - 6 -methoxy-2 -methylpyrimidine was found.
  • the present invention provides a compound of formula
  • each of R a and ⁇ t independently represents a C ⁇ -Cg alkoxy group or a halogen atom; a pharmaceutically acceptable metabolically labile ester or amide thereof; or a pharmaceutically acceptable salt of said compound, said ester or said amide.
  • the compound 2-hydroxymethyl-4-chloro-5- (imidazolidin- 2-ylidene) -6-methoxypyrimidine has been found to lower blood pressure and heart rate when administered intravenously to spontaneously hypertensive rats at a dose of 3 mg/kg when tested according to the method described by A. D. Palkowitz et al . , J. Med . Chem . , Vol. 37, p. 4508-4521, 1994.
  • the invention includes each tautomeric form of the compound .
  • Pharmaceutically acceptable salts of the compound of formula I include acid addition salts formed with inorganic acids such as hydrochloric, hydrobromic, hydriodic, sulfuric and phosphoric acid, as well as organic acids such as para- toluenesulfonic, methanesulfonic, oxalic, para- bromophenylsulfonic, carbonic, succinic, citric, benzoic, and acetic acid, and related inorganic and organic acids.
  • the pharmaceutically acceptable salts thus include the hydrogen chloride, hydrogen bromide and hydrogen iodide acid addition salts.
  • metabolically labile ester and amides of the compound of formula I are ester or amide derivatives of the compound of formula I that are hydrolyzed in vivo to afford the compound of formula I and a pharmaceutically acceptable carboxylic acid.
  • labile esters include esters formed with the hydroxyl group in the compound of formula I and an alkanoic acid, such as acetic acid.
  • metabolically labile amides include amides formed with an NH group in the compound of formula I and an alkanoic acid such as acetic acid.
  • a particular value for a C ] _-C alkoxy group represented by R a or Rb is methoxy.
  • a particular value for a halogen atom represented by R a or Rb is chlorine.
  • R a represents methoxy and R-b represents chlorine .
  • a particularly preferred compound is 2-hydroxymethyl-4- chloro-5- (imidazolidin-2-ylidene) -6-methoxypyrimidine .
  • the compound 2-hydroxymethyl-4-chloro-5- (imidazolidin- 2-ylidene) -6-methoxypyrimidine was originally isolated as a metabolite of moxonidine from the urine of rats and dogs.
  • Another aspect of the invention is, therefore, a compound of formula I in a pharmaceutically acceptable state of purity.
  • the compounds according to the invention may also be prepared by a process which comprises:
  • R 1 represents a hydrogen atom or a hydroxyl protecting group and R 2 represents a hydrogen atom or an amino protecting group;
  • R ⁇ represents a carboxyl group or an ester derivative thereof, or a salt thereof;
  • hydroxyl protecting groups include acyl groups, such as groups of formula R 4 CO in which
  • R 4 represents C_-Cg alkyl, (3-10C) cycloalkyl, phenyl C ⁇ -Cg alkyl, phenyl, C ] _-Cg alkoxy, phenyl C ⁇ -Cg alkoxy, or (3 -IOC) cycloalkoxy, wherein a phenyl group may be unsubstituted or substituted by one or two substituents independently selected from amino, hydroxy, nitro, halogeno, C ] _-Cg alkyl,
  • amine protecting groups include acyl groups of formula R 5 C0, in which R 5 is as defined for R 4 .
  • R 1 and R 2 are hydrogen and C ] _-Cg alkanoyl such as acetyl .
  • R ⁇ are carboxy and C ⁇ -Cg alkoxycarbonyl such as methoxycarbonyl or ethoxycarbonyl .
  • process step (a) the compound of formula II is conveniently reacted with the compound of formula III in the presence of a dehydrating agent, such as phosphorous oxychloride.
  • a dehydrating agent such as phosphorous oxychloride.
  • the reaction is conveniently conducted at a temperature in the range of from 20 to 150°C, preferably at 90 to 110°C.
  • An excess of phosphorous oxychloride may be used as reaction solvent.
  • the reduction process step (b) may conveniently be performed using a reducing agent such as lithium aluminum hydride. Suitable solvents include ethers such as tetrahydrofuran .
  • the base is conveniently an alkali metal alkoxide, such as a sodium alkoxide or potassium alkoxide.
  • the reaction is conveniently performed in an alkanol as a reaction solvent, and under reflux conditions.
  • Any protecting groups may conveniently be removed employing a method well known in the art.
  • an acyl group protecting the hydroxyl or an amino group may conveniently be removed by acid or base catalyzed hydrolysis or methanolysis, for example by reaction with methanol in the presence of an alkali metal methoxide .
  • an acyl group protecting the hydroxyl or an amino group in a compound of formula la may be removed during the replacement of a halogen atom with an alkoxy group.
  • metabolically labile esters or amides of the compound of formula I, and pharmaceutically acceptable salts thereof may be prepared by methods well known in the art .
  • the compounds of formula IV may be prepared by reacting a compound of formula
  • the compounds of formula II and V may be prepared by reducing a compound of formula
  • R 6 represents R 1 OCH2 or R 3 , as appropriate.
  • the reduction is conveniently performed using hydrogen and a group VIII metal catalyst, for example Raney nickel.
  • the compounds of formula VI may be prepared by reacting a compound of formula
  • a halogenating agent for example a phosphorus oxyhalide.
  • the reaction is conveniently performed in the presence of a base, such as N,N-diethylaniline .
  • a base such as N,N-diethylaniline .
  • One or two atoms may subsequently be replaced with alkoxy by reaction with an alkali metal alkoxide in an alkanol .
  • the compounds of formula VII may be prepared by reacting a compound of formula
  • nitrating agent for example a mixture of fuming nitric acid and glacial acetic acid.
  • the compounds of formula VIII may be prepared by reacting a compound of formula
  • a salt thereof such as the hydrochloride with a malonate, such as diethylmalonate, in the presence of a strong base, such as an alkali metal alkoxide, for example sodium ethoxide .
  • the compounds of formula III and IX are known, for example from US 4,323,570, or may be prepared by well-known methods. Certain intermediates described herein, for example the compounds of formula IV are believed to be novel and are provided as further aspects of the invention.
  • the compounds according to the invention are preferably formulated in a pharmaceutical composition prior to administration .
  • the present invention provides a compound of formula I, a pharmaceutically acceptable metabolically labile ester or amide thereof, or a pharmaceutically acceptable salt thereof, in combination with a pharmaceutically acceptable diluent or carrier.
  • compositions may be prepared by known procedures using well-known and readily available ingredients.
  • the active ingredient will usually be mixed with a carrier, or diluted by a carrier, or enclosed within a carrier, and may be in the form of a capsule, sachet, paper, or other container.
  • the carrier serves as a diluent, it may be a solid, semi-solid, or liquid material which acts as a vehicle, excipient, or medium for the active ingredient.
  • compositions can be in the form of tablets, pills, powders, lozenges, sachets, cachets, elixirs, suspensions, emulsions, solutions, syrups, aerosols, ointments containing, for example, up to 10% by weight of active compound, soft and hard gelatin capsules, suppositories, sterile injectable solutions, and sterile packaged powders .
  • Suitable carriers, excipients, and diluents include lactose, dextrose, sucrose, sorbitol, mannitol, starches, gum, acacia, calcium phosphate, alginates, tragacanth, gelatin, calcium silicate, micro- crystalline cellulose, polyvinylpyrrolidone, cellulose, water syrup, methyl cellulose, methyl and propyl hydroxy- benzoates, talc, magnesium stearate, and mineral oil.
  • the formulations can additionally include lubricating agents, wetting agents, emulsifying and suspending agents, preserving agents, sweetening agents, or flavoring agents.
  • Compositions of the invention may be formulated so as to provide quick, sustained, or delayed release of the active ingredient after administration to the patient by employing procedures well known in the art.
  • compositions are preferably formulated in a unit dosage form, each dosage containing from about 5 mg to about 500 mg, more preferably about 25 mg to about 300 mg of the active ingredient.
  • unit dosage form refers to a physically discrete unit suitable as unitary dosages for human subjects and other mammals, each unit containing a predetermined quantity of active material calculated to produce the desired therapeutic effect, in association with a suitable pharmaceutical carrier, diluent, or excipient .
  • suitable pharmaceutical carrier diluent, or excipient .
  • Tablets each containing 60 mg of active ingredient are made as follows :
  • the active ingredient, starch, and cellulose are passed through a No. 45 mesh U.S. sieve and mixed thoroughly.
  • the solution of polyvinylpyrrolidone is mixed with the resultant powders which are then passed through a No . 14 mesh U.S. sieve.
  • the granules so produced are dried at 50°C and passed through a No. 18 mesh U.S. sieve.
  • the sodium carboxymethyl starch, magnesium stearate, and talc previously passed through a No. 60 mesh U.S. sieve, are then added to the granules which, after mixing, are compressed on a tablet machine to yield tablets each weighing 150 mg.
  • the compounds according to the invention are useful in the treatment of hypertension, congestive heart failure, non-insulin dependent diabetes mellitus, smoking cessation, nicotine withdrawal, opioid withdrawal, ethanol withdrawal and atherosclerosis. According to other aspects of the invention therefore, the present invention provides the use of the compounds of the invention for the treatment of each of the conditions in a warm blooded mammal, such as a human.
  • a warm blooded mammal such as a human.
  • the following example illustrates the invention.
  • Spontaneously hypertensive rats (Tac :N (SHR) fBR) were obtained from Taconic Farms (Germantown, NY) at 14-16 weeks of age (295-325 gm) and housed under a 12-hour light/dark cycle (lights on from 0600 to 1800 hours) . Following a 1- week acclimation period, rats were anesthetized with 2% isoflurane (Aerrane, Anaquest, Madison, WI) for implantation of blood pressure transmitters (model TA11PA-C40, Data
  • a bulldog clamp was placed caudal to the left renal artery, and the aorta was punctured rostral to the common iliac using a 21-gauge needle (bent at a 45° angle with the bevel down) .
  • a fluid- filled catheter (0.7 mm OD, 8 cm in length) attached to the hermetically sealed transmitter was inserted and advanced to the bulldog clamp using the bent needle as a guide. The area was dried with a cotton swab and tissue adhesive applied at the entry point while the clamp was removed. The entry point was further sealed using tissue adhesive and a cellulose fiber patch (Vetland, 3M Co) .
  • the body of the transmitter was sutured to the muscles of the inner abdominal wall using non-absorbable 4-0 silk, the muscle layers were approximated with sterile 3-0 silk, and the final incision closed with sterile metal wound clips. All animals were administered 10,000 units of penicillin intramuscularly (Ambi-Pen®, Butler) , housed individually in shoe box cages with food and water ad libitum, and permitted to recover for at least one week before study. Digitized pressure signals were acquired for 30 seconds every 10 minutes using DSI Dataquest IV 2.0 software. Mean pressure was calculated as the arithmetic mean of the sample waveform sampled at a frequency of 500 Hz. The digitized values were stored and manipulated on a Compac Deskpro 486/33MHz computer.
  • Test compounds or vehicle were administered via the tail vein in a total volume of 0.3-0.4 ml.
  • the hydroxymethyl metabolite was administered in a water vehicle made by adding approximately 50 ⁇ l of IN HCL to the metabolite base and slowly diluting with water to appropriate concentration. Rats were then returned to telemetry cages and blood pressure was monitored for the subsequent 24 hours.
  • the hydroxymethyl metabolite lowered mean blood pressure compared to vehicle in the SHR.
  • the moxonidine hydroxymethyl metabolite has been identified in urine of rats, dogs, mice and humans.
  • Moxonidine and the hydroxymethyl metabolite are shown to be analagous in that they both possess antihypertensive and bradycardic activity in conscious unrestrained hypertensive rats .
  • This analogy can also be drawn from moxonidine to the hydroxymethyl metabolite in other treatments, as well, as would be apparent to those skilled in the art .
  • Moxonidine has been shown to be an effective treatment for hypertension (U.S. Patent No. 4,323,570), smoking cessation, nicotine withdrawal, opioid withdrawal, ethanol withdrawal (U.S. Patent No. 5,732,717), congestive heart failure (PCT publication W097/46241) , non-insulin dependent diabetes (U.S. Patent No. 5,712,283), and atherosclerosis (PCT publication W096/26728) . Therefore, another embodiment of this invention is the use of a compound of the formula
  • each of R a and R ⁇ independently represents a C ] _-Cg alkoxy group or a halogen atom; a pharmaceutically acceptable metabolically labile ester or amide thereof; or a pharmaceutically acceptable salt of said compound, said ester or said amide for the treatment of hypertension, smoking cessation, nicotine withdrawal, opioid withdrawal, ethanol withdrawal, congestive heart failure, non-insulin dependent diabetes, and atherosclerosis.
  • the particular dose of compound administered according to this invention for the treatment of the indications alone or in combination will of course be determined by the particular circumstances surrounding the case, including the compound administered, the warm blooded mammal being treated, the route of administration, the particular condition being treated, and similar considerations.
  • the compounds can be administered by a variety of routes including oral, rectal, transdermal , subcutaneous, intravenous, intramuscular, or intranasal routes. Alternatively, the compound may be administered by continuous infusion.
  • a typical daily dose will contain from 0.005 mg/kg to 50 mg/kg of the active compound of this invention.
  • daily doses will be 0.01 mg/kg to 25 mg/kg, more preferably from 0.1 mg/kg to 10 mg/kg.
  • Example 1 2-Hydroxymethyl-4-chloro-5- (imidazolin-2-ylideneimino) -6- methoxypyrimidine
  • Step (a) (2.3 g, 16.2 mmol) over the period of 40 min at 10- 15°C.
  • the reaction mixture was stirred for 5.5 hours at room temperature then cooled down to 5°C and diluted with cold water (3 mL) .
  • the precipitate was collected by filtration, washed with ethanol (5 L) and ether (5 mL) , and dried under vacuum to give 2.44 g (80.5%) of title compound as a colorless solid.
  • IR (KBr) 539, 789, 1105, 1297, 1358,

Landscapes

  • Chemical & Material Sciences (AREA)
  • Organic Chemistry (AREA)
  • Pharmaceuticals Containing Other Organic And Inorganic Compounds (AREA)

Abstract

The present invention relates to novel pyrimidine derivatives, to processes for preparing them, to intermediates useful in preparing them, to pharmaceutical compositions containing them and to uses for them as pharmaceuticals.

Description

Pyrimidine Derivatives
This application claims the benefit of U.S. Provisional Application No. 60/057,472, filed September 3, 1997. The present invention relates to novel pyrimidine derivatives, to processes for preparing them, to intermediates useful in preparing them, to pharmaceutical compositions containing them and to uses for them as pharmaceuticals . United States patent number 4,323,570 discloses a group of aminopyrimidine derivatives having antihypertensive activity. One of the compounds, 4-chloro-5- (imidazoline-2- ylamino) -6-methoxy-2-methylpyrimidine, is commercially available in some countries under the generic name moxonidine for the treatment of hypertension. This compound may interconvert with its tautomer, 4-chloro-5- (imidazolidin-2 -ylideneimino) - 6 -methoxy-2 -methylpyrimidine . We have now found a novel pyrimidine derivative having a 2-hydroxymethyl substituent which possesses anti- hypertensive activity.
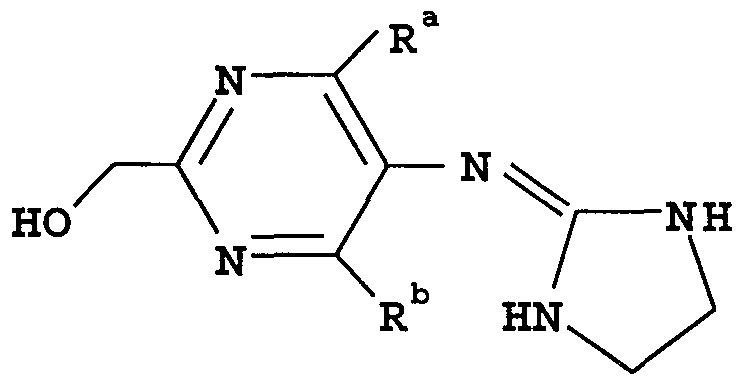
The present invention provides a compound of formula
Figure imgf000003_0001
in which each of Ra and ϊt independently represents a C^-Cg alkoxy group or a halogen atom; a pharmaceutically acceptable metabolically labile ester or amide thereof; or a pharmaceutically acceptable salt of said compound, said ester or said amide.
The compound 2-hydroxymethyl-4-chloro-5- (imidazolidin- 2-ylidene) -6-methoxypyrimidine has been found to lower blood pressure and heart rate when administered intravenously to spontaneously hypertensive rats at a dose of 3 mg/kg when tested according to the method described by A. D. Palkowitz et al . , J. Med . Chem . , Vol. 37, p. 4508-4521, 1994.
It will be appreciated by those skilled in the art that the compounds of formula I can exist in tautomeric form as shown below.
Figure imgf000004_0001
The invention includes each tautomeric form of the compound .
Pharmaceutically acceptable salts of the compound of formula I include acid addition salts formed with inorganic acids such as hydrochloric, hydrobromic, hydriodic, sulfuric and phosphoric acid, as well as organic acids such as para- toluenesulfonic, methanesulfonic, oxalic, para- bromophenylsulfonic, carbonic, succinic, citric, benzoic, and acetic acid, and related inorganic and organic acids. The pharmaceutically acceptable salts thus include the hydrogen chloride, hydrogen bromide and hydrogen iodide acid addition salts.
Pharmaceutically acceptable metabolically labile ester and amides of the compound of formula I are ester or amide derivatives of the compound of formula I that are hydrolyzed in vivo to afford the compound of formula I and a pharmaceutically acceptable carboxylic acid. Examples of labile esters include esters formed with the hydroxyl group in the compound of formula I and an alkanoic acid, such as acetic acid. Example of metabolically labile amides include amides formed with an NH group in the compound of formula I and an alkanoic acid such as acetic acid.
A particular value for a C]_-C alkoxy group represented by Ra or Rb is methoxy.
A particular value for a halogen atom represented by Ra or Rb is chlorine.
Preferably Ra represents methoxy and R-b represents chlorine .
A particularly preferred compound is 2-hydroxymethyl-4- chloro-5- (imidazolidin-2-ylidene) -6-methoxypyrimidine . The compound 2-hydroxymethyl-4-chloro-5- (imidazolidin- 2-ylidene) -6-methoxypyrimidine was originally isolated as a metabolite of moxonidine from the urine of rats and dogs. Another aspect of the invention is, therefore, a compound of formula I in a pharmaceutically acceptable state of purity. The compounds according to the invention may also be prepared by a process which comprises:
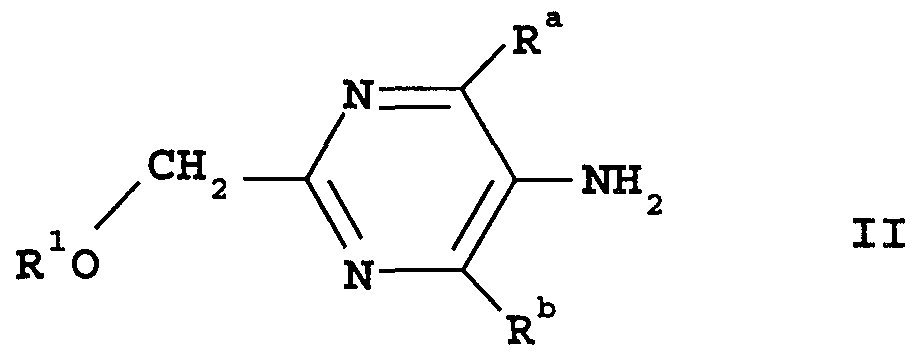
(a) reacting a compound of formula
Figure imgf000005_0001
or a salt thereof, with a compound of formula
Figure imgf000005_0002
in which R1 represents a hydrogen atom or a hydroxyl protecting group and R2 represents a hydrogen atom or an amino protecting group; or
(b) reducing a compound of formula
Figure imgf000006_0001
in which R^ represents a carboxyl group or an ester derivative thereof, or a salt thereof; to afford a compound of formula
Figure imgf000006_0002
of a salt thereof; whereafter, as necessary, optionally
(i) replacing a halogen atom represented by Ra or R)° with a C]_-Cg alkoxy group by reaction with a C]_-Cg alkanol in the presence of a base; and further optionally
(ii) removing any remaining protecting groups; and further optionally
(iii) forming a pharmaceutically acceptable metabolically labile ester or amide; or
(iv) forming a pharmaceutically acceptable salt. The protection of hydroxyl and amino groups, is generally described in McOmie, Protecting Groups in Organic Chemistry, Plenum Press, NY, 1973, and Greene and Wuts, Protecting Groups in Organic Synthesis, 2nd. Ed., John Wiley & Sons, NY, 1991. Examples of hydroxyl protecting groups include acyl groups, such as groups of formula R4CO in which
R4 represents C_-Cg alkyl, (3-10C) cycloalkyl, phenyl C^-Cg alkyl, phenyl, C]_-Cg alkoxy, phenyl C^-Cg alkoxy, or (3 -IOC) cycloalkoxy, wherein a phenyl group may be unsubstituted or substituted by one or two substituents independently selected from amino, hydroxy, nitro, halogeno, C]_-Cg alkyl,
C]_-Cg alkoxy, carboxy, C^-Cg alkoxycarbonyl , carbamoyl ,
C]_-Cg alkanoylamino, C]_-Cg alkylsulphonylamino, phenyl- sulphonylamino, toluenesulphonylamino, and C^-Cg fluoro- alkyl . Examples of amine protecting groups include acyl groups of formula R5C0, in which R5 is as defined for R4.
Examples of particular values for R1 and R2 are hydrogen and C]_-Cg alkanoyl such as acetyl . Examples of particular values for R^ are carboxy and C^-Cg alkoxycarbonyl such as methoxycarbonyl or ethoxycarbonyl .
In process step (a) , the compound of formula II is conveniently reacted with the compound of formula III in the presence of a dehydrating agent, such as phosphorous oxychloride. The reaction is conveniently conducted at a temperature in the range of from 20 to 150°C, preferably at 90 to 110°C. An excess of phosphorous oxychloride may be used as reaction solvent. The reduction process step (b) may conveniently be performed using a reducing agent such as lithium aluminum hydride. Suitable solvents include ethers such as tetrahydrofuran .
In the replacement of a halogen atom with an alkoxy group, the base is conveniently an alkali metal alkoxide, such as a sodium alkoxide or potassium alkoxide. The reaction is conveniently performed in an alkanol as a reaction solvent, and under reflux conditions. Any protecting groups may conveniently be removed employing a method well known in the art. For example, an acyl group protecting the hydroxyl or an amino group may conveniently be removed by acid or base catalyzed hydrolysis or methanolysis, for example by reaction with methanol in the presence of an alkali metal methoxide . In this connection, it will be appreciated that an acyl group protecting the hydroxyl or an amino group in a compound of formula la may be removed during the replacement of a halogen atom with an alkoxy group.
Pharmaceutically acceptable metabolically labile esters or amides of the compound of formula I, and pharmaceutically acceptable salts thereof may be prepared by methods well known in the art . The compounds of formula IV may be prepared by reacting a compound of formula
Figure imgf000008_0001
or a salt thereof with a compound of formula III, following the method of process step (a) .
The compounds of formula II and V may be prepared by reducing a compound of formula
Figure imgf000008_0002
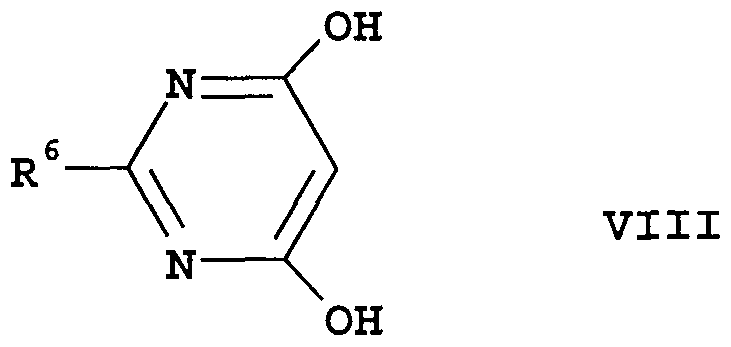
in which R6 represents R1OCH2 or R3 , as appropriate. The reduction is conveniently performed using hydrogen and a group VIII metal catalyst, for example Raney nickel. The compounds of formula VI may be prepared by reacting a compound of formula
Figure imgf000009_0001
with a halogenating agent, for example a phosphorus oxyhalide. The reaction is conveniently performed in the presence of a base, such as N,N-diethylaniline . One or two atoms may subsequently be replaced with alkoxy by reaction with an alkali metal alkoxide in an alkanol .
The compounds of formula VII may be prepared by reacting a compound of formula
Figure imgf000009_0002
with a nitrating agent, for example a mixture of fuming nitric acid and glacial acetic acid.
The compounds of formula VIII may be prepared by reacting a compound of formula
NH
R6— C
\ IX
NH2
or a salt thereof, such as the hydrochloride with a malonate, such as diethylmalonate, in the presence of a strong base, such as an alkali metal alkoxide, for example sodium ethoxide .
The compounds of formula III and IX are known, for example from US 4,323,570, or may be prepared by well-known methods. Certain intermediates described herein, for example the compounds of formula IV are believed to be novel and are provided as further aspects of the invention.
The compounds according to the invention are preferably formulated in a pharmaceutical composition prior to administration .
According to another aspect, therefore, the present invention provides a compound of formula I, a pharmaceutically acceptable metabolically labile ester or amide thereof, or a pharmaceutically acceptable salt thereof, in combination with a pharmaceutically acceptable diluent or carrier.
The pharmaceutical compositions may be prepared by known procedures using well-known and readily available ingredients. In making the compositions of the present invention, the active ingredient will usually be mixed with a carrier, or diluted by a carrier, or enclosed within a carrier, and may be in the form of a capsule, sachet, paper, or other container. When the carrier serves as a diluent, it may be a solid, semi-solid, or liquid material which acts as a vehicle, excipient, or medium for the active ingredient. The compositions can be in the form of tablets, pills, powders, lozenges, sachets, cachets, elixirs, suspensions, emulsions, solutions, syrups, aerosols, ointments containing, for example, up to 10% by weight of active compound, soft and hard gelatin capsules, suppositories, sterile injectable solutions, and sterile packaged powders .
Some examples of suitable carriers, excipients, and diluents include lactose, dextrose, sucrose, sorbitol, mannitol, starches, gum, acacia, calcium phosphate, alginates, tragacanth, gelatin, calcium silicate, micro- crystalline cellulose, polyvinylpyrrolidone, cellulose, water syrup, methyl cellulose, methyl and propyl hydroxy- benzoates, talc, magnesium stearate, and mineral oil. The formulations can additionally include lubricating agents, wetting agents, emulsifying and suspending agents, preserving agents, sweetening agents, or flavoring agents. Compositions of the invention may be formulated so as to provide quick, sustained, or delayed release of the active ingredient after administration to the patient by employing procedures well known in the art.
The compositions are preferably formulated in a unit dosage form, each dosage containing from about 5 mg to about 500 mg, more preferably about 25 mg to about 300 mg of the active ingredient. The term "unit dosage form" refers to a physically discrete unit suitable as unitary dosages for human subjects and other mammals, each unit containing a predetermined quantity of active material calculated to produce the desired therapeutic effect, in association with a suitable pharmaceutical carrier, diluent, or excipient . The following formulation example is illustrative only and is not intended to limit the scope of the invention in any way.
Formulation Example
Tablets each containing 60 mg of active ingredient are made as follows :
Active Ingredient 60 mg
Starch 45 mg
Microcrystalline cellulose 35 mg
Polyvinylpyrrolidone 4 mg
Sodium carboxymethyl starch 4.5 mg Magnesium stearate 0.5 mg
Talc 1 mg
Total 150 mg
The active ingredient, starch, and cellulose are passed through a No. 45 mesh U.S. sieve and mixed thoroughly. The solution of polyvinylpyrrolidone is mixed with the resultant powders which are then passed through a No . 14 mesh U.S. sieve. The granules so produced are dried at 50°C and passed through a No. 18 mesh U.S. sieve. The sodium carboxymethyl starch, magnesium stearate, and talc, previously passed through a No. 60 mesh U.S. sieve, are then added to the granules which, after mixing, are compressed on a tablet machine to yield tablets each weighing 150 mg.
The compounds according to the invention are useful in the treatment of hypertension, congestive heart failure, non-insulin dependent diabetes mellitus, smoking cessation, nicotine withdrawal, opioid withdrawal, ethanol withdrawal and atherosclerosis. According to other aspects of the invention therefore, the present invention provides the use of the compounds of the invention for the treatment of each of the conditions in a warm blooded mammal, such as a human. The following example illustrates the invention.
Example Biological Activity in Hypertensive Rats
Telemetry Implantation
Spontaneously hypertensive rats (Tac :N (SHR) fBR) were obtained from Taconic Farms (Germantown, NY) at 14-16 weeks of age (295-325 gm) and housed under a 12-hour light/dark cycle (lights on from 0600 to 1800 hours) . Following a 1- week acclimation period, rats were anesthetized with 2% isoflurane (Aerrane, Anaquest, Madison, WI) for implantation of blood pressure transmitters (model TA11PA-C40, Data
Sciences Int (DSI) , St Paul, MN) as follows: The abdomen was shaved, scrubbed with Betadine®, and a 4.5-cm abdominal incision was made beginning just caudal to the approximate location of the kidneys. The abdominal aorta was isolated and gently cleaned of connective tissue with a sterile cotton swab. A small spatula was used to raise a portion of the aorta away from the vena cava in an area just rostral to -li¬
the iliac artery bifurcation. A bulldog clamp was placed caudal to the left renal artery, and the aorta was punctured rostral to the common iliac using a 21-gauge needle (bent at a 45° angle with the bevel down) . A fluid- filled catheter (0.7 mm OD, 8 cm in length) attached to the hermetically sealed transmitter was inserted and advanced to the bulldog clamp using the bent needle as a guide. The area was dried with a cotton swab and tissue adhesive applied at the entry point while the clamp was removed. The entry point was further sealed using tissue adhesive and a cellulose fiber patch (Vetland, 3M Co) . The body of the transmitter was sutured to the muscles of the inner abdominal wall using non-absorbable 4-0 silk, the muscle layers were approximated with sterile 3-0 silk, and the final incision closed with sterile metal wound clips. All animals were administered 10,000 units of penicillin intramuscularly (Ambi-Pen®, Butler) , housed individually in shoe box cages with food and water ad libitum, and permitted to recover for at least one week before study. Digitized pressure signals were acquired for 30 seconds every 10 minutes using DSI Dataquest IV 2.0 software. Mean pressure was calculated as the arithmetic mean of the sample waveform sampled at a frequency of 500 Hz. The digitized values were stored and manipulated on a Compac Deskpro 486/33MHz computer.
Protocol
Animals were briefly sedated with 2% isoflurane. Test compounds or vehicle were administered via the tail vein in a total volume of 0.3-0.4 ml. The hydroxymethyl metabolite was administered in a water vehicle made by adding approximately 50 μl of IN HCL to the metabolite base and slowly diluting with water to appropriate concentration. Rats were then returned to telemetry cages and blood pressure was monitored for the subsequent 24 hours.
Monitoring was initiated immediately except that the values for 15 min prior and subsequent to dosing were omitted. Results
The hydroxymethyl metabolite lowered mean blood pressure compared to vehicle in the SHR.
Table 1
Figure imgf000014_0001
The effect of the hydroxymethyl metabolite lasted for approximately 4 hours. Accompanying the effect of hydroxymethyl metabolite on blood pressure was pronounced falls in heart rate (Table 2) that were most prominent 4 hours after dosing.
Table 2
Figure imgf000014_0002
Figure imgf000015_0001
Consistent with the shorter effect of the hydroxymethyl metabolite on blood pressure, there was also a significant fall compared to vehicle in heart rate apparent during the first hour after dosing.
Conclusion
The moxonidine hydroxymethyl metabolite has been identified in urine of rats, dogs, mice and humans.
Moxonidine and the hydroxymethyl metabolite are shown to be analagous in that they both possess antihypertensive and bradycardic activity in conscious unrestrained hypertensive rats . This analogy can also be drawn from moxonidine to the hydroxymethyl metabolite in other treatments, as well, as would be apparent to those skilled in the art . Moxonidine has been shown to be an effective treatment for hypertension (U.S. Patent No. 4,323,570), smoking cessation, nicotine withdrawal, opioid withdrawal, ethanol withdrawal (U.S. Patent No. 5,732,717), congestive heart failure (PCT publication W097/46241) , non-insulin dependent diabetes (U.S. Patent No. 5,712,283), and atherosclerosis (PCT publication W096/26728) . Therefore, another embodiment of this invention is the use of a compound of the formula
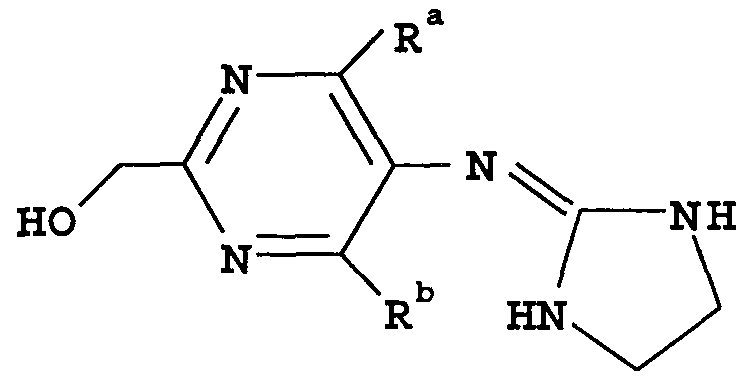
Figure imgf000016_0001
in which each of Ra and R^ independently represents a C]_-Cg alkoxy group or a halogen atom; a pharmaceutically acceptable metabolically labile ester or amide thereof; or a pharmaceutically acceptable salt of said compound, said ester or said amide for the treatment of hypertension, smoking cessation, nicotine withdrawal, opioid withdrawal, ethanol withdrawal, congestive heart failure, non-insulin dependent diabetes, and atherosclerosis.
The particular dose of compound administered according to this invention for the treatment of the indications alone or in combination will of course be determined by the particular circumstances surrounding the case, including the compound administered, the warm blooded mammal being treated, the route of administration, the particular condition being treated, and similar considerations. The compounds can be administered by a variety of routes including oral, rectal, transdermal , subcutaneous, intravenous, intramuscular, or intranasal routes. Alternatively, the compound may be administered by continuous infusion. A typical daily dose will contain from 0.005 mg/kg to 50 mg/kg of the active compound of this invention. Preferably, daily doses will be 0.01 mg/kg to 25 mg/kg, more preferably from 0.1 mg/kg to 10 mg/kg.
The following Example illustrates the invention. Example 1 2-Hydroxymethyl-4-chloro-5- (imidazolin-2-ylideneimino) -6- methoxypyrimidine
(a) Preparation of 4 , 6-dihydroxy-2-hydroxymethylpyrimidine : To a solution of sodium ethoxide prepared from sodium (1.38 g, 60.0 mmol) and ethanol (29 mL) , was added hydroxyacetamidine hydrochloride (McCasland, G. E. and Tarbell, S. - ". Amer . Chem. Soc . , 1946, 68, 2393-2395) (2.21 g, 20.0 mmol) and after 5 min ethyl malonate (3.2 g, 20.0 mmol) . The reaction mixture was refluxed for 3 hours then cooled to room temperature, diluted with water (10 L) and acidified with concentrated hydrochloric acid (4 mL) . The precipitate was filtered off, washed with water (10 mL) , ethanol (5 mL) and ether (5 mL) , and dried under vacuum to give 2.24 g (79%) of the title compound as a light-brown solid. IR (KBr) : 527, 1106, 1324, 1571, 1639, 1682, 2910,
3029, 3084 cm-1-, UV (EtOH) λ max(ε): 256 (4235), 239 nm
(3398); 1H-NMR (DMSO-dg) δ 4.28(s), 5.14(s), 5.66(s); MS(FD): 142 (M+, 100%).
(b) Preparation of 4 , 6-dihydroxy-2-hydroxymethyl-5- nitropyrimidine :
To a stirred mixture of nitric acid (fuming, 4.2 mL) and acetic acid (glacial, 2.1 mL) was added the product of
Step (a) (2.3 g, 16.2 mmol) over the period of 40 min at 10- 15°C. The reaction mixture was stirred for 5.5 hours at room temperature then cooled down to 5°C and diluted with cold water (3 mL) . The precipitate was collected by filtration, washed with ethanol (5 L) and ether (5 mL) , and dried under vacuum to give 2.44 g (80.5%) of title compound as a colorless solid. IR (KBr) : 539, 789, 1105, 1297, 1358,
1630, 1674, 2828, 2920, 3402, 3460 cm"1-, UV (EtOH) λ max(ε) : 322 (3354), 213 nm (17626); XH-NMR (DMSO-dg) δ 4.42 (s,CH2); MS (FD) : 187 (M+, 100%). (c) Preparation of 2-acetoxymethyl-4 , 6-dihydroxy-5- nitropyrimidine :
A mixture of the product of Step (b) (2.35 g, 12.56 mmol), acetic acid (glacial, 15 mL) , and acetic anhydride (15 mL) was heated at 105°C for 2 hours, then evaporated under vacuum, and re-evaporated with toluene (3 times) . The residue was dried under vacuum to give 2.843 g (99%) of title compound as a yellow solid. IR (KBr) : 545, 787, 1212, 1326, 1354, 1367, 1493, 1650, 1673, 1758, 2939, 3082, 3141 cm"1-, UV (EtOH) λ max(ε) : 327 nm (3365); iH-NMR (DMS0-d6) δ
2.14 (s, 3H, CH3), 4.98 (s, 2H, CH2); MS (FD) : 229 (M+, 100%) .
(d) Preparation of 2-acetoxymethyl-4 , 6-dichloro-5- nitropyrimidine :
To a mixture of the product of Step (c) (2.75 g, 12.0 mmol) and phosphorus oxychloride (13 mL) was added N,N- diethylaniline (2.5 mL, 15.7 mmol) dropwise. The reaction mixture was refluxed for 2 hours and evaporated under vacuum. The residue was diluted with ether (15 mL) and poured onto ice. After stirring for 3 min organic layer was separated, washed with saturated aqueous solutions of sodium bicarbonate and sodium chloride, then dried over sodium sulfate and evaporated under vacuum. Flash chromatography of the residue on silica gel (20% ether in hexane) gave 2.446 g (77%) of title compound as a white solid, Rf 0.60
(hexane/ethyl acetate, 3:2). IR (KBr): 836, 853, 1218,
1550, 1755 cm-1-, UV (EtOH) λ max(ε): 259 nm (3290); !H-NMR (CDC13) δ 2.28 (s, 3H, CH3 ) , 5.33 (s , 2H, CH2). Analysis calculated for C7H5Cl2N3θ4: C, 31.60; H, 1.89; N, 15.79.
Found: C, 31.84; H, 1.85; N, 15.69. (e) Preparation of 2-acetoxymethyl-4 , 6-dichloro-5- aminopyrimidine :
A mixture of the product of Step (d) (255 mg, 0.96 mmol) and Raney nickel (30 mg) in ethanol (25 mL) was hydrogenated in Parr apparatus (50 pSi of hydrogen) for 5.5 h. The resulting suspension was filtered through celite and evaporated under vacuum. Flash chromatography of the residue on silica gel (40% ethyl acetate in hexane) gave 202 mg (89%) of title compound as a colorless solid, Rf 0.41 (hexane/ethyl acetate, 3:2). IR (KBr): 793, 907, 1249,
1366, 1459, 1513, 1621, 1731, 3332, 3428 cm-1-, UV (EtOH) λ max(ε) : 314 (5676), 253 nm (11629); ^-H-NMR (CDC13) δ 2.21 (s, 3H, CH3), 4.55 (s, 2H, NH2 ) , 5.16 (s, 2H, CH2 ) ; MS (FD) :
235 (M+, 100%) .
(f) Preparation of 2-acetoxymethyl-4 , 6-dichloro-5- (1- acetylimidazolidin-2-ylidenimino) pyrimidine :
To a solution of the product of Step (e) (191 mg, 0.81 mmol) in phosphorus oxychloride (1 mL) was added N-acetyl-2- imidazolidone (114 mg, 0.89 mmol) in one portion. The reaction mixture was heated at 105-110°C (bath) for 3 hours and evaporated under vacuum. The residue was diluted with ice water (2 mL) , made alkaline with aqueous sodium hydroxide ( 5N, 2 mL) and extracted with dichloromethane (3 x 5 mL) . The extract was washed with saturated aqueous solutions of sodium bicarbonate and sodium chloride, dried over sodium sulfate and evaporated under vacuum. Flash chromatography of the residue on silica gel (from 50 to 60% of ethyl acetate in hexane) gave 166 mg (59%) of title compound as a colorless solid, Rf 0.21 (hexane/ethyl acetate, 1:1). IR (KBr): 815, 1033, 1074, 1219, 1355, 1378, 1414, 1497, 1657, 1675, 1752, 3254, cm"1-, UV (EtOH) λ max(ε) : 245 nm (12118); iH-ΝMR (CDCI3) δ 2.23 (s, 3H, CH3COO) , 2.70 (s, 3H, CH3COΝ) , 3.58 (t, J = 7.98 Hz, 2H, CH2N) , 4.11 (t, J = 7.98 Hz, 2H, CH2N) , 4.74 (s, 1H, NH) , 5.19 (s, 2H, CH20) ; HRMS (FAB): calculated for C12H13C12N503 : 346.0474. Found: 346.0481.
(g) Preparation of 2-hydroxymethyl-4-chloro-5- (imidazolidin-2-ylidenimino) -6-methoxypyrimidine
A solution of the product of Step (f) (87 mg, 0.25 mmol) and sodium methoxide (14 mg, 0.26 mmol) in methanol (1.2 mL) was refluxed for 3 hours. The reaction mixture was evaporated under vacuum to one half of the initial volume and cooled to 5°C. The precipitate was collected by filtration, washed with water (1 mL) and dried under vacuum to give 32 mg (50%) of title compound as a colorless solid. IR (KBr): 720, 1042, 1091, 1283, 1307, 1376, 1440, 1471, 1533, 1659, 3030, 3225, 3340 cm"1-, UV (EtOH) λ max(ε): 254 nm (8577); i-H-NMR (DMSO-dg) δ 3.34 (s, 4H, CH2N) , 3.87 (s,
3H, CH3), 4.39 (d, J = 6.18 Hz, 2H, CH20) , 5.19 (t, J = 6.18
Hz, 1H, OH), 6.26 (s, 2H, NH) , HRMS (FAB): calculated for C9H12C1N502: 258.0758. Found: 258.0759.

Claims

We Claim :
1. A compound of formula
Figure imgf000021_0001
in which each of Ra and R^ independently is a C]_-Cg alkoxy group or a halogen atom; a pharmaceutically acceptable metabolically labile ester or amide thereof; or a pharmaceutically acceptable salt of said compound, said ester or said amide.
2. A compound as claimed in Claim 1, in which Ra is methoxy and R^3 is chlorine.
3. The compound of Claim 1, which is 2 -hydroxymethyl -4- chloro-5- (imidazolin-2-ylideneimino) -6-methoxypyrimidine .
4. A compound as claimed in any one of Claims 1 to 3 , which is in a pharmaceutically acceptable state of purity.
5. A pharmaceutical composition, which comprises a compound as claimed in Claim 1 together with a pharmaceutically acceptable diluent or carrier.
6. A method of treating hypertension, congestive heart failure, non-insulin dependent diabetes, smoking cessation, nicotine withdrawal, opioid withdrawal, ethanol withdrawal, or atherosclerosis in a warm blooded mammal requiring such treatment, which comprises administering an effective amount of a compound as claimed in Claim 1.
7. A process for preparing a compound as claimed in Claim 1, which comprises:
(a) reacting a compound of formula
Figure imgf000022_0001
or a salt thereof, with a compound of formula
R2
in which R1 is a hydrogen atom or a hydroxyl protecting group and R2 is a hydrogen atom or an amino protecting group; or
(b) reducing a compound of formula
Figure imgf000022_0003
in which R3 is a carboxyl group or an ester derivative thereof, or a salt thereof; to afford a compound of formula
Figure imgf000023_0001
of a salt thereof; and optionally,
(i) replacing a halogen atom of Ra or R3 with a C^-Cg alkoxy group by reaction with C^-Cg alkanol in the presence of a base; and optionally
(ii) removing any remaining protecting groups; and optionally
(iii) forming a pharmaceutically acceptable metabolically labile ester or amide; and optionally
(iv) forming a pharmaceutically acceptable salt.
8. A compound of formula
Figure imgf000023_0002
in which Ra and R5 each independently are a C -Cg alkoxy group or a halogen atom, R2 is a hydrogen atom or an amino protecting group, and R3 is a carboxyl group or an esterified carboxyl group, or a salt thereof.
9. A compound as claimed in Claim 10, in which R2 is a hydrogen atom or a C1-C6 alkanoyl group and R3 represents a carboxyl group or a C^-Cg alkoxycarbonyl group.
10. A compound of formula
Figure imgf000024_0001
in which each of Ra and Rb independently represents a (1- 6C) alkoxy group or a halogen atom; a pharmaceutically acceptable metabolically labile ester or amide thereof; or a pharmaceutically acceptable salt of said compound, said ester or said amide.
11. A compound as claimed in Claim 10, in which Ra is methoxy and Rb is chlorine.
12. The compound of Claim 10, which is 2 -hydroxymethyl-4- chloro-5- (imidazolin-2-ylideneimino) -6-methoxypyrimidine .
13. A compound as claimed in any one of Claims 10 to 12, which is in a pharmaceutically acceptable state of purity.
14. A pharmaceutical composition, which comprises a compound as claimed in Claim 10 together with a pharmaceutically acceptable diluent or carrier.
15. A compound as claimed in Claim 10 for use in therapy.
16. The use of a compound as claimed in Claim 10 for the manufacture of a medicament for the treatment of hypertension, congestive heart failure, non-insulin dependent diabetes, smoking cessation, nicotine withdrawal, opioid withdrawal, ethanol withdrawal or atherosclerosis.
17. A process for preparing a compound as claimed in Claim 10, which comprises:
(a) reacting a compound of formula
Figure imgf000025_0001
or a salt thereof, with a compound of formula
R2
Figure imgf000025_0002
in which R1 represents a hydrogen atom or a hydroxyl protecting group and R2 represents a hydrogen atom or an amino protecting group; or
(b) reducing a compound of formula
Figure imgf000025_0003
in which R3 represents a carboxyl group or an ester derivative thereof, or a salt thereof; to afford a compound of formula
Figure imgf000026_0001
of a salt thereof; whereafter, as necessary,
(i) replacing a halogen atom represented by Ra or Rb with a (1-6C) alkoxy group by reaction with (1-6C) alkanol in the presence of a base;
(ii) removing any remaining protecting groups;
(iii) forming a pharmaceutically acceptable metabolically labile ester or amide; and
(iv) forming a pharmaceutically acceptable salt.
18. A compound of formula
Figure imgf000026_0002
in which Ra and Rb each independently represent a (1- 6C) alkoxy group or a halogen atom, R2 represents a hydrogen atom or an amino protecting group, and R3 represents a carboxyl group or an esterified carboxyl group, or a salt thereof .
19. A compound as claimed in Claim 18, in which R2 represents a hydrogen atom or a C^ alkanoyl group and R3 represents a carboxyl group or a (1-6C) alkoxycarbonyl group.
0 . A compound of formula
Figure imgf000027_0001
in which each of Ra and Rb independently represents a (1- 6C) alkoxy group or a halogen atom; a pharmaceutically acceptable metabolically labile ester or amide thereof; or a pharmaceutically acceptable salt of said compound, said ester or said amide.
21. A compound as claimed in Claim 20, in which Ra is methoxy and Rb is chlorine.
22. The compound of Claim 20, which is 2 -hydroxymethyl-4- chloro-5- (imidazolin-2-ylideneimino) -6-methoxypyrimidine .
23. A compound as claimed in any one of Claims 20 to 22, which is in a pharmaceutically acceptable state of purity.
24. A pharmaceutical formulation adapted for the treatment of hypertension, congestive heart failure, non-insulin dependent diabetes, smoking cessation, opioid withdrawal, ethanol withdrawal or atherosclerosis comprising as an active ingredient a compound having the Formula
Figure imgf000028_0001
in which each of Ra and Rb independently represents a (1- 6C) alkoxy group or a halogen atom; a pharmaceutically acceptable metabolically labile ester or amide thereof; or a pharmaceutically acceptable salt of said compound, said ester or said amide.
25. The formulation of Claim 24 adapted for the treatment of hypertension, congestive heart failure, non-insulin dependent diabetes, smoking cessation, nicotine withdrawal, opioid withdrawal, ethanol withdrawal or atherosclerosis comprising as an active ingredient a compound having the Formula comprising as an active ingredient a compound of Formula I wherein which Ra is methoxy and Rb is chlorine.
26. The formulation of Claim 24 adapted for the treatment of hypertension, congestive heart failure, non- insulin dependent diabetes, smoking cessation, opioid withdrawal, ethanol withdrawal or atherosclerosis comprising as an active ingredient a compound having the Formula comprising as an active ingredient a compound of Formula I wherein the compound is 2-hydroxymethyl-4-chloro-5- (imidazolin-2-ylideneimino) -6-methoxypyrimidine .
PCT/US1998/018381 1997-09-03 1998-09-03 Pyrimidine derivatives Ceased WO1999011269A1 (en)

Priority Applications (3)

Application Number Priority Date Filing Date Title
AU93022/98A AU9302298A (en) 1997-09-03 1998-09-03 Pyrimidine derivatives
CA002269971A CA2269971A1 (en) 1997-09-03 1998-09-03 Pyrimidine derivatives
EP98945876A EP0954316A4 (en) 1997-09-03 1998-09-03 Pyrimidine derivatives

Applications Claiming Priority (2)

Application Number Priority Date Filing Date Title
US5747297P 1997-09-03 1997-09-03
US60/057,472 1997-09-03

Publications (1)

Publication Number Publication Date
WO1999011269A1 true WO1999011269A1 (en) 1999-03-11

Family

ID=22010758

Family Applications (1)

Application Number Title Priority Date Filing Date
PCT/US1998/018381 Ceased WO1999011269A1 (en) 1997-09-03 1998-09-03 Pyrimidine derivatives

Country Status (4)

Country Link
EP (1) EP0954316A4 (en)
AU (1) AU9302298A (en)
CA (1) CA2269971A1 (en)
WO (1) WO1999011269A1 (en)

Citations (1)

* Cited by examiner, † Cited by third party
Publication number Priority date Publication date Assignee Title
US4323570A (en) * 1978-11-15 1982-04-06 Beiersdorf Aktiengesellschaft Substituted aminopyrimidines

Family Cites Families (1)

* Cited by examiner, † Cited by third party
Publication number Priority date Publication date Assignee Title
MA18829A1 (en) * 1979-05-18 1980-12-31 Ciba Geigy Ag PYRIMIDINE DERIVATIVES, METHODS FOR THEIR PREPARATION, PHARMACEUTICAL COMPOSITIONS CONTAINING THESE COMPOUNDS AND THEIR THERAPEUTIC USE

Patent Citations (1)

* Cited by examiner, † Cited by third party
Publication number Priority date Publication date Assignee Title
US4323570A (en) * 1978-11-15 1982-04-06 Beiersdorf Aktiengesellschaft Substituted aminopyrimidines

Non-Patent Citations (1)

* Cited by examiner, † Cited by third party
Title
See also references of EP0954316A4 *

Also Published As

Publication number Publication date
AU9302298A (en) 1999-03-22
EP0954316A1 (en) 1999-11-10
CA2269971A1 (en) 1999-03-11
EP0954316A4 (en) 2002-02-13

Similar Documents

Publication Publication Date Title
JP3447292B2 (en) Imidazopyridines and their use for the treatment of gastrointestinal diseases
BE1006343A5 (en) Derivatives indole.
RU2067975C1 (en) 1,2-dihydro-2-oxopyridines, their physiologically acceptable salts and pharmaceutical composition
US5332750A (en) 1,2-dihydro-2-oxopyridines
CZ180195A3 (en) Azetinediones being substituted by spirocycloalkyl group, their use and process of their preparation, pharmaceutical composition containing such substituted azetinediones, process of its preparation and a composition in which said preparation is comprised
ZA200301863B (en) Xanthine phosphodiesterase V inhibitors.
JPWO1991001306A1 (en) Oxindole derivatives
JPS6364428B2 (en)
JP7283832B2 (en) Pyridine sulfonamide phosphate compound or its pharmaceutically acceptable salt, its production method, its use and pharmaceutical composition
CN101570510B (en) Quinoline compound, pharmaceutical composition, preparation method and application thereof
FR2687676A1 (en) NOVEL ANTAGONIST POLYAZAINDEN DERIVATIVES OF ANGIOTENSIN II RECEPTORS; PROCESSES FOR PREPARING THEM, PHARMACEUTICAL COMPOSITIONS CONTAINING SAME
JPH0249306B2 (en)
EP0529654A1 (en) Benzopyran derivatives as potassium channel activators and 5-lipoxygenase inhibitors
JP5683276B2 (en) Method for producing bosentan
WO1998033763A1 (en) Substituted hydroxy-anilino derivatives of cyclobutene-3,4-diones
US20030144321A1 (en) Antitumor agents
JP5290190B2 (en) Salt of imidazole-5-carboxylic acid derivative, production method and pharmaceutical composition thereof
KR920005827B1 (en) Benzothiadiazepine derivatives
KR20030078962A (en) Alkoxycarbonylamino heteroaryl carboxylic acid derivatives as ip antagonists
EP0954316A1 (en) Pyrimidine derivatives
JPH0665209A (en) Imidazolyl-substituted phenylpropionic acid and cinnamic acid derivatives
JPH07291936A (en) Novel imidazole derivative and method for producing the same
US5578596A (en) 2-alkoxy-5,6,7,8-tetrahydroquinoxaline derivatives, and production method and use thereof
JPS6130588A (en) Benzo(c)(1,8)naphthylidine, manufacture, use and medicine
JP2837318B2 (en) Angiotensin II antagonistic pyridine derivative

Legal Events

Date Code Title Description
AK Designated states

Kind code of ref document: A1

Designated state(s): AL AM AT AU AZ BA BB BG BR BY CA CH CN CU CZ DE DK EE ES FI GB GE GH GM HR HU ID IL IS JP KE KG KP KR KZ LC LK LR LS LT LU LV MD MG MK MN MW MX NO NZ PL PT RO RU SD SE SG SI SK SL TJ TM TR TT UA UG US UZ VN YU ZW

AL Designated countries for regional patents

Kind code of ref document: A1

Designated state(s): GH GM KE LS MW SD SZ UG ZW AM AZ BY KG KZ MD RU TJ TM AT BE CH CY DE DK ES FI FR GB GR IE IT LU MC NL PT SE BF BJ CF CG CI CM GA GN GW ML MR NE SN TD TG

WWE Wipo information: entry into national phase

Ref document number: 1998945876

Country of ref document: EP

ENP Entry into the national phase

Ref document number: 2269971

Country of ref document: CA

Ref country code: CA

Ref document number: 2269971

Kind code of ref document: A

Format of ref document f/p: F

121 Ep: the epo has been informed by wipo that ep was designated in this application
WWP Wipo information: published in national office

Ref document number: 1998945876

Country of ref document: EP

REG Reference to national code

Ref country code: DE

Ref legal event code: 8642

NENP Non-entry into the national phase

Ref country code: JP

Ref document number: 1999517042

Format of ref document f/p: F

WWW Wipo information: withdrawn in national office

Ref document number: 1998945876

Country of ref document: EP