WO1999043680A1 - Substituted pyrrolobenzimidazoles for treating inflammatory diseases - Google Patents

Substituted pyrrolobenzimidazoles for treating inflammatory diseases Download PDF

Info

Publication number
WO1999043680A1
WO1999043680A1 PCT/US1999/004085 US9904085W WO9943680A1 WO 1999043680 A1 WO1999043680 A1 WO 1999043680A1 US 9904085 W US9904085 W US 9904085W WO 9943680 A1 WO9943680 A1 WO 9943680A1
Authority
WO
WIPO (PCT)
Prior art keywords
compound
pyridyl
fluorophenyl
pyrrolo
dihydro
Prior art date
Legal status (The legal status is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the status listed.)
Ceased
Application number
PCT/US1999/004085
Other languages
French (fr)
Inventor
John H. Dodd
James A. Henry
Kenneth C. Rupert
Current Assignee (The listed assignees may be inaccurate. Google has not performed a legal analysis and makes no representation or warranty as to the accuracy of the list.)
Janssen Pharmaceuticals Inc
Original Assignee
Ortho McNeil Pharmaceutical Inc
Priority date (The priority date is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the date listed.)
Filing date
Publication date
Application filed by Ortho McNeil Pharmaceutical Inc filed Critical Ortho McNeil Pharmaceutical Inc
Priority to JP2000533434A priority Critical patent/JP4570777B2/en
Priority to DE69917005T priority patent/DE69917005T2/en
Priority to EP99908456A priority patent/EP1060181B1/en
Priority to CA002321559A priority patent/CA2321559A1/en
Priority to AU27888/99A priority patent/AU759297B2/en
Priority to AT99908456T priority patent/ATE266029T1/en
Publication of WO1999043680A1 publication Critical patent/WO1999043680A1/en
Anticipated expiration legal-status Critical
Ceased legal-status Critical Current

Links

Classifications

    • C—CHEMISTRY; METALLURGY
    • C07—ORGANIC CHEMISTRY
    • C07D—HETEROCYCLIC COMPOUNDS
    • C07D487/00—Heterocyclic compounds containing nitrogen atoms as the only ring hetero atoms in the condensed system, not provided for by groups C07D451/00 - C07D477/00
    • C07D487/02—Heterocyclic compounds containing nitrogen atoms as the only ring hetero atoms in the condensed system, not provided for by groups C07D451/00 - C07D477/00 in which the condensed system contains two hetero rings
    • C07D487/04—Ortho-condensed systems
    • A—HUMAN NECESSITIES
    • A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
    • A61P1/00—Drugs for disorders of the alimentary tract or the digestive system
    • A61P1/04—Drugs for disorders of the alimentary tract or the digestive system for ulcers, gastritis or reflux esophagitis, e.g. antacids, inhibitors of acid secretion, mucosal protectants
    • A—HUMAN NECESSITIES
    • A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
    • A61P1/00—Drugs for disorders of the alimentary tract or the digestive system
    • A61P1/16—Drugs for disorders of the alimentary tract or the digestive system for liver or gallbladder disorders, e.g. hepatoprotective agents, cholagogues, litholytics
    • A—HUMAN NECESSITIES
    • A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
    • A61P1/00—Drugs for disorders of the alimentary tract or the digestive system
    • A61P1/18—Drugs for disorders of the alimentary tract or the digestive system for pancreatic disorders, e.g. pancreatic enzymes
    • A—HUMAN NECESSITIES
    • A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
    • A61P13/00—Drugs for disorders of the urinary system
    • A61P13/12—Drugs for disorders of the urinary system of the kidneys
    • A—HUMAN NECESSITIES
    • A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
    • A61P17/00—Drugs for dermatological disorders
    • A61P17/06—Antipsoriatics
    • A—HUMAN NECESSITIES
    • A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
    • A61P19/00—Drugs for skeletal disorders
    • A61P19/02—Drugs for skeletal disorders for joint disorders, e.g. arthritis, arthrosis
    • A—HUMAN NECESSITIES
    • A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
    • A61P19/00—Drugs for skeletal disorders
    • A61P19/08—Drugs for skeletal disorders for bone diseases, e.g. rachitism, Paget's disease
    • A61P19/10—Drugs for skeletal disorders for bone diseases, e.g. rachitism, Paget's disease for osteoporosis
    • A—HUMAN NECESSITIES
    • A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
    • A61P21/00—Drugs for disorders of the muscular or neuromuscular system
    • A61P21/04—Drugs for disorders of the muscular or neuromuscular system for myasthenia gravis
    • A—HUMAN NECESSITIES
    • A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
    • A61P25/00—Drugs for disorders of the nervous system
    • A—HUMAN NECESSITIES
    • A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
    • A61P29/00—Non-central analgesic, antipyretic or antiinflammatory agents, e.g. antirheumatic agents; Non-steroidal antiinflammatory drugs [NSAID]
    • A—HUMAN NECESSITIES
    • A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
    • A61P3/00—Drugs for disorders of the metabolism
    • A61P3/08—Drugs for disorders of the metabolism for glucose homeostasis
    • A61P3/10—Drugs for disorders of the metabolism for glucose homeostasis for hyperglycaemia, e.g. antidiabetics
    • A—HUMAN NECESSITIES
    • A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
    • A61P37/00—Drugs for immunological or allergic disorders
    • A61P37/02—Immunomodulators
    • A61P37/06—Immunosuppressants, e.g. drugs for graft rejection
    • A—HUMAN NECESSITIES
    • A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
    • A61P37/00—Drugs for immunological or allergic disorders
    • A61P37/08—Antiallergic agents
    • A—HUMAN NECESSITIES
    • A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
    • A61P9/00—Drugs for disorders of the cardiovascular system
    • A61P9/02—Non-specific cardiovascular stimulants, e.g. drugs for syncope, antihypotensives

Definitions

  • This invention relates to substituted pyrrolobenzimidazoles, and related pharmaceutical compositions and methods for treating inflammatory diseases.
  • the compounds of the invention inhibit the production of cytokines, particularly TNF- ⁇ and IL-1, which mediate inflammatory responses.
  • Rheumatoid arthritis is an inflammatory disease which affects millions of people and can affect any joint in the human body. Its symptoms range from mild pain and inflammation in affected joints, to severe and debilitating pain and inflammation. Although the disease is associated mainly with aging adults, it is not restricted to adults.
  • NSAID nonsteroidal anti-inflammatory drugs
  • drugs such as methotrexate, gold salts, D-penicillamine and prednisone are often used when patients fail to respond to NSAID's. These drugs also have significant toxicities and their mechanisms of action remain unknown.
  • SB 203580 Due to SB 203580' s oral activity and potency in animal models, researchers have suggested that a compound with this profile has potential as a viable treatment for rheumatoid arthritis.
  • a and B are independently nitrogen or carbon, at least one of A and B being nitrogen;
  • D and E are nitrogen, with the proviso that (i) a double bond exists between the non-aryl carbon and either D or E, (ii) R 2 is absent if the double bond exists between the non-aryl carbon and D, and (iii) R 3 is absent if the double bond exists between the non-aryl carbon and E; and
  • subject means any animal or artificially modified animal having an inflammatory disease.
  • the subject is a human.
  • Second Embodiment of Formula I Scheme 3 can be used to prepare the compound of Formula I, wherein A is carbon, ring 1 has a 4-chloro substituent, and B is nitrogen.
  • the hydrazinobenzimida- zole 3_a is coupled with the 2-phenyl-l-pyridyl substituted ethanone, 3b, in a suitable solvent such as ethylene glycol at about 90 °C for about 1-3 hours to give the illustrated compound.
  • Similar compounds of Formula I may be prepared by varying the ethanone and benzimidazole starting materials.
  • the ethanone starting material, 3b is prepared from the addition, hydrolysis and subsequent decarboxylation of a phenylacetonitrile derivative and methylisonicotinate.
  • % inhibition [1- (sample -BKG) / (CTRL-BKG) ] x 100.
  • Compound 1 inhibited 44% of the activity of p38 at 20 ⁇ M.
  • the majority of the mononuclear cell layer was carefully removed using a pipet and these PBMC s were diluted with PBS and spun at 600 x g for 15 minutes. The resulting PBMC s were washed with another portion of PBS and spun at 400 x g for 10 minutes at room temperature.
  • the recovered pellets were diluted in low endotoxin RPMI / 1% FCS culture medium and gave a cell concentration of 0.5-2.0 x 10⁶ PBMC/ mL. A small volume of the suspension was removed for counting on a hemocytometer and the remaining preparation was centrifuged at 200 x g for 15 minutes at room temperature. The recovered pelleted PBM s were resuspended in RPMI / 1% FCS to a concentration of 1.67 x 10⁶/mL.
  • the PBMC suspension (180 ⁇ L) was transferred to duplicate wells of a 96-well flat-bottom microtiter plate and incubated for 1 hour at 37 °C.
  • a solution of test compound (10 ⁇ L: prepared at 20 x the desired final concentration) was added to each well and the plate was incubated for 1 hour at 37 °C.
  • a solution (10 ⁇ L) of LPS in RPMI / 1% FCS (200 ng/mL) was added and the wells were incubated overnight at 37°C.
  • the supernatant 100 ⁇ L was removed from each well and diluted with RPMI / 1% FCS (400 ⁇ L) .
  • the samples were analyzed for TNF- ⁇ using a commercial ELISA kit (Genzyme) .
  • the anti IL-l ⁇ activity of certain compounds of the invention was determined by the following in vitro assay.
  • Plastic-adherent cells were prepared for PBMC. Briefly, PBMCs were added to the wells of a 96-well plate as above, incubated for lh at 37 °C, and the adherent cells prepared by gently re-suspending the non-adherent cells with a pipettor, removing and discarding the supernatant, and gently washing the wells 3 times with 200 ⁇ L of culture medium. Additional culture medium (180 ⁇ L) was added to the wells after the final wash. Compound addition, LPS stimulation, incubation and supernatant harvest were as for TNF- ⁇ .
  • mice BALB / cJ females, Jackson Laboratories
  • rats Lewis males, Charles River
  • mice were fasted for 30 minutes prior to oral dosing with 5-10 mL/kg of test compound at 5-50 mg/kg.
  • the animals were injected intraperitoneally with LPS at 1 mg/kg and returned to their cages for 1 hour. Animals were anesthetized by C0 , exsanguinated by cardiac puncture and whole blood collected (0.1-0.7 mL) .

Landscapes

  • Health & Medical Sciences (AREA)
  • Chemical & Material Sciences (AREA)
  • Organic Chemistry (AREA)
  • Engineering & Computer Science (AREA)
  • Bioinformatics & Cheminformatics (AREA)
  • Life Sciences & Earth Sciences (AREA)
  • Public Health (AREA)
  • General Chemical & Material Sciences (AREA)
  • Medicinal Chemistry (AREA)
  • Nuclear Medicine, Radiotherapy & Molecular Imaging (AREA)
  • Veterinary Medicine (AREA)
  • Pharmacology & Pharmacy (AREA)
  • Chemical Kinetics & Catalysis (AREA)
  • Animal Behavior & Ethology (AREA)
  • General Health & Medical Sciences (AREA)
  • Immunology (AREA)
  • Physical Education & Sports Medicine (AREA)
  • Rheumatology (AREA)
  • Orthopedic Medicine & Surgery (AREA)
  • Diabetes (AREA)
  • Cardiology (AREA)
  • Neurology (AREA)
  • Heart & Thoracic Surgery (AREA)
  • Pain & Pain Management (AREA)
  • Urology & Nephrology (AREA)
  • Biomedical Technology (AREA)
  • Hematology (AREA)
  • Neurosurgery (AREA)
  • Endocrinology (AREA)
  • Emergency Medicine (AREA)
  • Obesity (AREA)
  • Transplantation (AREA)
  • Pulmonology (AREA)
  • Gastroenterology & Hepatology (AREA)
  • Dermatology (AREA)
  • Pharmaceuticals Containing Other Organic And Inorganic Compounds (AREA)
  • Nitrogen Condensed Heterocyclic Rings (AREA)

Abstract

This invention provides compounds, and pharmaceutically acceptable salts thereof, having structure (I), as well as related pharmaceutical compositions, and methods of treatment and synthesis. A and B are C or N, R1-R3 are as in the application.

Description

SUBSTITUTED PYRROLOBENZIMIDAZOLES FOR TREATING INFLAMMATORY DISEASES
Throughout this application, various publications are cited. The disclosure of these publications is hereby incorporated by reference into this application to describe more fully the state of the art to which this invention pertains.
Field of the Invention
This invention relates to substituted pyrrolobenzimidazoles, and related pharmaceutical compositions and methods for treating inflammatory diseases. The compounds of the invention inhibit the production of cytokines, particularly TNF-α and IL-1, which mediate inflammatory responses.
Background of the Invention
The inflammatory cytokines IL-1 and TNF-α play an important role in a number of inflammatory diseases. (C. Dinarello et al . , Inflammatory cytokines: Interleukin-1 and Tumor Necrosis Factor as Effector Molecules in Autoimmune Diseases, Curr. Opin . Immunol . 1991, 3, 941- 48.) Rheumatoid arthritis is a prime example of such inflammatory diseases, and is thus the inflammatory disease focused on most in this section.
Rheumatoid arthritis is an inflammatory disease which affects millions of people and can affect any joint in the human body. Its symptoms range from mild pain and inflammation in affected joints, to severe and debilitating pain and inflammation. Although the disease is associated mainly with aging adults, it is not restricted to adults.
The most common rheumatoid arthritis therapy involves the use of nonsteroidal anti-inflammatory drugs (NSAID's) to alleviate symptoms. However, despite the widespread use of NSAID's, many individuals cannot tolerate the doses necessary to treat the disease over a prolonged period of time. In addition, NSAID's merely treat the symptoms of disease without affecting the underlying cause (s). Other drugs, such as methotrexate, gold salts, D-penicillamine and prednisone are often used when patients fail to respond to NSAID's. These drugs also have significant toxicities and their mechanisms of action remain unknown.
Receptor antagonists to IL-1 and monoclonal antibodies to TNF-α have been shown to reduce symptoms of rheumatoid arthritis in small-scale human clinical trials. (M.J. Elliot et al., Treatment of Rheumatoid Arthritis with Chimeric Monoclonal Antibodies to Tumor
Necrosis Factor α, Arthri tis Rheum. 1993 36, 1681-90.)
In addition to protein-based therapies, there are small molecule agents which inhibit the production of these cytokines and have demonstrated activity in animal rheumatoid arthritis models. (J.C. Boehm et al . , 1- Substituted 4-Aryl-5-pyridinylimidazoles: A New Class of Cytokine Suppressive Drugs with Low 5-Lipoxygenase and Cyclooxygenase Inhibitory Potency, J. Med. Chem. , 1996, 39, 3929-37.) Of these small molecule agents, SB 203580 has proven effective in reducing the production of TNF-α and IL-1 in LPS-sti ulated human monocyte cell lines with IC50 values of 50 to 100 nM. (J. Adams et al., Imidazole Derivatives And Their Use as Cytokine Inhibitor, International Patent Application WO 93/14081, 1993.)
Figure imgf000005_0001
SB 203580
In addition to this in vitro behavior, SB 203580 has been shown to inhibit the production of inflammatory cytokines in rats and mice at IC5o values of 15 to 25 mg/kg. (A.M. Badger, et al., Pharmacological Profile of SB 203580, A Selective Inhibitor of Cytokine Suppressive Binding Protein/p38 Kinase, in Animal Models of Arthritis, Bone Resorption, Endotoxin Shock and Immune Function, The Journal of Pharmacology and Experimental Therapeutics, 1996, 219, 1453-61.)
Due to SB 203580' s oral activity and potency in animal models, researchers have suggested that a compound with this profile has potential as a viable treatment for rheumatoid arthritis. (A.M. Badger, et al . Pharmacological Profile of SB 203580, A Selective Inhibitor of Cytokine Suppressive Binding Protein/p38 Kinase, in Animal Models of Arthritis, Bone Resorption, Endotoxin Shock and Immune Function, The Journal of
Pharmacology and Experimental Therapeutics, 1996, 279, 1453-61.)
SB 203580 and other small molecules reduce the production of inflammatory cytokines by inhibiting the activity of a serine/threonine kinase, p38 (also referred to in the art as "CSBP") , at an IC50 of 200 nM. (D. Griswold et al., Pharmacology of Cytokine Suppressive Anti-Inflammatory Drug Binding Protein (CSPB) , A Novel Stress-Induced Kinase, Pharmacology Communications, 1996, 7, 323-29.) Although the precise mechanism of this kinase is unknown, it has been implicated in both the production of TNF-α and the signaling responses associated with the TNF-α receptor.
Rheumatoid arthritis, and the host of other inflammatory disorders, take a severe toll on those afflicted. There is thus a tremendous need for small molecule anti-inflammatory agents. To date, however, no such agent — including SB 203580 — has ever been shown to be anti-inflammatory in human clinical trials.
Summary of the Invention
This invention provides a compound having the structure
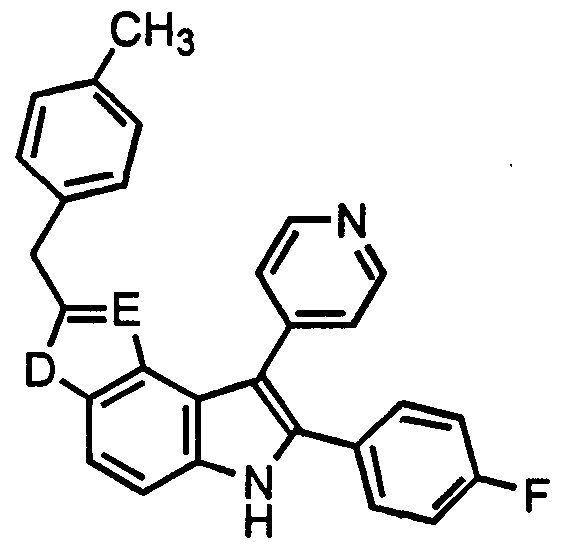
Figure imgf000007_0001
or a pharmaceutically acceptable salt thereof, wherein:
(a) Ri, R2 and R3 are independently selected from the group consisting of (i) hydrogen, (ii) Cι-5alkyl, (iii) Cι-5alkylamino, (iv) diCι-5alkylamino, (v) a phenyl substituted with one or more of hydrogen, halogen, Ci-salkyl, and trihaloCi-salkyl, and (vi) a phenylCι-5alkyl substituted with one or more of hydrogen, halogen, Cι_5alkyl, and trihaloCι-5alkyl;
(b) rings 1 and 2 are each independently substituted with one or more substituents selected from the group consisting of hydrogen, halogen, Cι-5alkyl, and trihaloCι-5alkyl;
(c) A and B are independently nitrogen or carbon, at least one of A and B being nitrogen; (d) D and E are nitrogen, with the proviso that (i) a double bond exists between the non-aryl carbon and either D or E, (ii) R2 is absent if the double bond exists between the non-aryl carbon and D, and (iii) R3 is absent if the double bond exists between the non-aryl carbon and E; and (e) the compound is neither 1, 6-dihydro-7- (4-pyridyl) -8- (4-fluorophenyl) -2-phenylmethyl-pyrrolo [3, 2- e]benzimidazole, nor 3, 6-dihydro-8- (4-fluorophenyl) - 3- (3-phenylpropyl) -7- (4-pyridyl) -pyrrolo [3,2- e]benzimidazole.
This invention also provides a pharmaceutical composition comprising the instant compound, and a pharmaceutically acceptable carrier. This invention further provides a method of treating a subject having an inflammatory disease, which comprises administering to the subject a therapeutically effective dose of the instant pharmaceutical composition.
Finally, this invention provides a method of making the instant compound, which comprises the step of contacting a first compound having the structure
Figure imgf000008_0001
with a second compound having the structure
Figure imgf000008_0002
under conditions permitting a Fischer indolization between the first and second compounds. Detailed Description of the Invention
This invention provides compounds and related pharmaceutical compositions and methods useful in the treatment of inflammatory diseases. The compounds of the invention inhibit the production of the inflammatory cytokines TNF-α and IL-lβ, the overproduction of which are characteristic of inflammatory diseases.
Specifically, this invention provides a compound having the structure
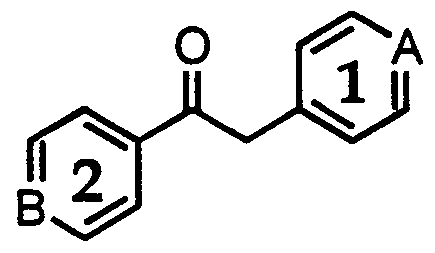
Figure imgf000009_0001
or a pharmaceutically acceptable salt thereof, wherein:
(a) Ri, R2 and R3 are independently selected from the group consisting of (i) hydrogen, (ii) Cι-5alkyl, (iii) Cι-5alkylamino, (iv) diCι-5alkylamino, (v) a phenyl substituted with one or more of hydrogen, halogen, Cι-5alkyl, and trihaloCι_5alkyl, and (vi) a phenylCi-δalkyl substituted with one or more of hydrogen, halogen, Cι_5alkyl, and trihaloCι-5alkyl;
(b) rings 1 and 2 are each independently substituted with one or more substituents selected from the group consisting of hydrogen, halogen, Cι-5alkyl, and trihaloCι-5alkyl;
(c) A and B are independently nitrogen or carbon, at least one of A and B being nitrogen; (d) D and E are nitrogen, with the proviso that (i) a double bond exists between the non-aryl carbon and either D or E, (ii) R2 is absent if the double bond exists between the non-aryl carbon and D, and (iii) R3 is absent if the double bond exists between the non-aryl carbon and E; and
(e) the compound is neither 1, 6-dihydro-7- (4-pyridyl) -8- (4-fluorophenyl) -2-phenylmethyl-pyrrolo [3,2- e]benzimidazole, nor 3, 6-dihydro-8- (4-fluorophenyl) - 3- (3-phenylpropyl) -7- (4-pyridyl) -pyrrolo [3, 2- e] benzimidazole .
In one embodiment of the instant compound, D and E are both nitrogen. In another embodiment, A is nitrogen and B is carbon. In still another embodiment, D, E and A are nitrogen and B is carbon. In the preferred embodiment of the instant compound, the compound is selected from the group consisting of (i) 1, 6-dihydro-7- (4-fluorophenyl) -8- (4-pyridyl) -2-phenyl-pyrrolo [3,2- e] benzimidazole; (ii) 1, 6-dihydro-7- (4-fluorophenyl) -8- (4-pyridyl) -2-butyl-pyrrolo [3, 2-e]benzimidazole; (iii) 1, 6-dihydro-7- (4-fluorophenyl) -8- (4-pyridyl) -2- (2- phenylethyl) -pyrrolo [3, 2-e] benzimidazole; (iv) 1,6- dihydro-7- (4-pyridyl) -8- (4-fluorophenyl) -pyrrolo [3,2- e] benzimidazole; and (v) 1, 6-dihydro-7- (4-fluorophenyl) - 8- (4-pyridyl) -2-phenylmethyl-pyrrolo [3, 2-e]benzimidazole .
As used herein, the terms below have the following meanings in relation to the instant compound: "independently", when in reference to chemical substituents, shall mean that when more than one substituent exists, the substituents may be the same or different; "alkyl" shall mean straight, cyclic and branched-chain alkyl; "alkoxy" shall mean O-alkyl; "halogen" shall mean fluorine, chlorine, bromine or iodine; and "Ph" shall mean phenyl. Typically the instant compound is isolated and used as a free base. However, its various embodiments can also be isolated and used as pharmaceutically acceptable salts. Examples of such salts include hydrobromic, hydroiodic, hydrochloric, perchloric, sulfuric, maleic, fumaric, malic, tartaric, citric, benzoic, andelic, methanesulfonic, hydroethanesulfonic, benzenesulfonic, oxalic, palmoic, 2- naphthalenesulfonic, p-toluenesulfonic, cyclohexanesulfamic and saccharic.
This invention also provides a pharmaceutical composition comprising the instant compound, and a pharmaceutically acceptable carrier.
Pharmaceutically acceptable carriers are well known to those skilled in the art and include, but are not limited to, 0.01-0.1 M and preferably 0.05 M phosphate buffer or 0.8% saline. Additionally, such pharmaceutically acceptable carriers can be aqueous or non-aqueous solutions, suspensions, and emulsions. Examples of non- aqueous solvents are propylene glycol, polyethylene glycol, vegetable oils such as olive oil, and injectable organic esters such as ethyl oleate. Aqueous carriers include water, ethanol, alcoholic/aqueous solutions, glycerol, emulsions or suspensions, including saline and buffered media. Oral carriers can be elixers, syrups, capsules tablets and the like. The typical solid carrier is an inert substance such as lactose, starch, glucose, methyl cellulose, magnesium sterate, dicalcium phosphate, mannitol and the like. Parenteral carriers include sodium chloride solution, Ringer's dextrose, dextrose and sodium chloride, lactated Ringer's and fixed oils. Intravenous carriers include fluid and nutrient replenishers, electrolyte replenishers such as those based on Ringer's dextrose, and the like. Preservatives and other additives can also be present, such as, for example, antimicrobials, antioxidants, chelating agents, inert gases and the like. All carriers can be mixed as needed with disintegrants, diluents, granulating agents, lubricants, binders and the like using conventional techniques known in the art.
This invention further provides a method of treating a subject having an inflammatory disease, which comprises administering to the subject a therapeutically effective dose of the instant pharmaceutical composition.
As used herein, "subject" means any animal or artificially modified animal having an inflammatory disease. In the preferred embodiment, the subject is a human.
Inflammatory diseases are numerous and well known in the art. Examples of inflammatory diseases include, but are in no way limited to, rheumatoid arthritis, insulin- dependent diabetes mellitus, multiple sclerosis, myasthenia gravis, Crohn' s disease, autoimmune nephritis, primary biliary cirrhosis, psoriasis, acute pancreatitis, allograph rejection, allergic inflammation, inflammatory bowel disease, septic shock, osteoporosis, osteoarthritis, and cognitive deficits induced by neuronal inflammation. In the preferred embodiment, the autoimmune disease is rheumatoid arthritis.
Administering the pharmaceutical composition can be effected or performed using any of the various methods known to those skilled in the art. The administering can be performed, for example, intravenously, intramuscularly, orally and subcutaneously. In the preferred embodiment, the instant pharmaceutical composition is administered orally. Additionally, the administering can comprise giving the subject a plurality of dosages over a suitable period of time which can be determined according to routine methods.
A "therapeutically effective dose" of the pharmaceutical composition means an amount sufficient to stop, reverse or reduce the progression of the inflammatory disease being treated. Methods are known in the art which can be used to determine therapeutically effective doses for administering the instant pharmaceutical composition in a subject. The effective dose for administering the pharmaceutical composition to a human, for example, would be determined mathematically from the results of animal studies. In one embodiment, the therapeutically effective dose is a dose sufficient to deliver from about .05 g to about 200 mg of the instant pharmaceutical composition per kilogram of body weight daily. In another embodiment, the therapeutically effective dose is a dose sufficient to deliver from about 0.5 mg to about 50 mg.
Finally, this invention provides a method of making the instant compound, which comprises the step of contacting a first compound having the structure
Figure imgf000014_0001
with a second compound having the structure
Figure imgf000014_0002
under conditions permitting a Fischer indolization between the first and second compounds. In the first and second compounds, the designations for the rings and ring substituents (i.e., rings 1 and 2, atoms A, B, D and E, and substituents Ri, R2 and R3) and definitions thereof, are the same as those used in connection with the instant compound described above. Conditions permitting Fischer indolizations are well known in the art, and are exemplified in the Experimental Details below.
This invention will be better understood by reference to the Experimental Details which follow, but those skilled in the art will readily appreciate that the specific experiments detailed are only illustrative of the invention as described more fully in the claims which follow thereafter. Experimental Details
A. Definitions
As used herein, the terms below shall have the following meanings in relation to the instant experimental methods: "FCS" shall mean fetal calf serum; "TCA" shall mean trichloroacetic acid; "RPMI" shall mean the medium from the Roswell Park Memorial Institute having Sigma Cat No. R0833; and "Formula I" shall mean the structure set forth below and defined above.
Figure imgf000015_0001
B. Syntheses The compounds of this invention can be prepared by the following schemes, whereby some schemes produce more than one of the instant compounds. In those cases, the choice of scheme is a matter of discretion which is well within the capabilities of those skilled in the art.
1. Pyrrolobenzimidazole Ring System As illustrated by Scheme 1, the pyrrolobenzimidazole ring system of compounds of Formula 1 is formed by coupling a 1, 2-substituted ethanone, lb, with an appropriately substituted 5-hydrazinobenzimidazole la, in a suitable solvent such as ethylene glycol at about 90 °C for about 1-3 hours to give the compound shown. Aside from the illustrated product, many other compounds of Formula I can be produced by varying the starting materials la and lb. A variety of substituted ethanones can be prepared by treating known benzamide derivatives with 4-picolyl anions. (Gallagher, T.F. et al., Regulation of Stress-Induced Cytokine Production by Pyridinylimidazoles; Inhibition of CSBP Kinase, Bioorganic & Medicinal Chemistry, 1997, 5, 49-64.) A variety of substituted hydrazinyl-benzimidazoles may be prepared by treating 5-amino benzimidazoles with nitrous acid and stannous chloride. (Chetverikov, V.P. et al., Synthesis of 2-imidazo [4, 5-e] indoles from 5- aminobenzimidazoles, Zhim. Geterotsiki . Soedin, 1980, 1 , 74-8.) Therefore, to prepare a compound of Formula I where A is nitrogen, B is carbon, ring 2 has a 4-fluoro substituent, and Ri is methylphenyl, replace the illustrated ^a with 2-methyl-5-hydrazinobenzimidazole and lb with 1- [4-fluorophenyl] -2- [4-pyridinyl] ethanone.
SCHEME 1
la
Figure imgf000016_0001
Figure imgf000016_0002
Figure imgf000016_0003
2. First Embodiment of Formula I Scheme 2 can be used to prepare the compound of Formula I, wherein A is nitrogen, B is carbon, R3 is ethyl, and Ri is hydrogen. The starting substituted ethanone, 2b, is treated with a suitable 6- hydrazinobenzimidazole 2a, in a suitable solvent such as ethylene glycol at about 90 °C for about 1-3 hours to give the desired compound. Aside from the illustrated compound other compounds of Formula I can be prepared in this manner. The 6-hydrazinobenzimidazoles can be prepared from the 6-amino compounds using similar methods as described in the preparation of the 5- hydrazinobenzimidazoles .
SCHEME 2
2a
Figure imgf000017_0001
2b
3. Second Embodiment of Formula I Scheme 3 can be used to prepare the compound of Formula I, wherein A is carbon, ring 1 has a 4-chloro substituent, and B is nitrogen. The hydrazinobenzimida- zole 3_a, is coupled with the 2-phenyl-l-pyridyl substituted ethanone, 3b, in a suitable solvent such as ethylene glycol at about 90 °C for about 1-3 hours to give the illustrated compound. Similar compounds of Formula I may be prepared by varying the ethanone and benzimidazole starting materials. The ethanone starting material, 3b, is prepared from the addition, hydrolysis and subsequent decarboxylation of a phenylacetonitrile derivative and methylisonicotinate. (Lantos, I. et al., Synthetic and Mechanistic Studies on the Preparation of Pyridyl- Substituted Imidazothiazoles, J. Org. Chem. , 1988, 53, 4223-27.)
SCHEME 3
3a
Figure imgf000018_0001
Figure imgf000018_0002
Figure imgf000018_0003
3b 4. 1, 6-Dihydro-7- (4-fluorophenyl) -8- (4-pyridyl) -2- phenylmethyl-pyrrolo [3,2-e] benzimidazole
Figure imgf000019_0001
2-Benzyl-5-hydrazinobenzimidazole dihydrochoride (7.62 g) and 1- (4-fluoro-phenyl) -2- (4-pyridinyl) ethanone (5.27 g) were dissolved in ethylene glycol (70 mL) . The mixture was heated and stirred at 90 °C for 1 hour, and at 160 °C for 3 hours. The resulting mixture was cooled to room temperature, poured into water (500 ml) , and neutralized with solid K2C03. The aqueous phase was extracted with ethyl acetate (2 x 600 ml) . The organic layers were separated, combined, washed with water (2 x 300 ml) and brine (200 ml), dried (Na2S04) and concentrated in vacuo ♦ The concentrated solution was filtered through an 8 x 2 inch plug of silica gel using 100% ethyl acetate (2 L) as eluent. Concentration in vacuo gave an off-white solid (2.65 g) . Recrystallization from ethanol gave a white solid: mp 167-69 °C; XH NMR
(300 MHz, DMSO-d6) : d 12.20 (IH, s) , 11.84 (IH, s) , 8.48 (2H, br s), 7.62 (2H, br s) , 7.45 (2H, br t) , 7.36-7.20 (9H, ) , 4.20 (2H, s) ; MS m/s MH+ 419; Anal. Calcd. for C27Hι9FN4-.38H2θ C 76.35, H 4.68, N 13.19. Found C 76.29, H 4.69, N 13.15. 1.38 wt % H20 found by Karl Fisher.
C. Assays
1. In Vitro Enzyme Assay
The biological activities of the compounds of this invention were demonstrated by in vitro assays. As discussed previously, agents which inhibit the activity of the enzyme p38 inhibit the production of the inflammatory cytokines TNF-α and IL-1. Compounds of the invention were measured for their ability to inhibit the activity of p38 by the following in vitro assay.
A solution (38 μL) of purified recombinant p38 (where the amount of enzyme was determined empirically considering the linear range of the assay and the acceptable signal to noise ratio; 6xHis-p38 expressed in E. coli) , myelin basic protein substrate (also determined empirically), and a buffer of pH 7.5 (Hepes:25 mM, MgCl2:10 mM, MnCl2:10 mM) were added to 92 wells of a 96- well round bottom polypropylene plate. The remaining wells were used for control ("CTRL") and background ("BKG") .
The CTRL was prepared with the enzyme, substrate buffer and 2% DMSO, and the BKG was prepared with substrate buffer and 2% DMSO. A solution (12 μL) of the test compound in DMSO (compounds were diluted to 125 μM in 10% DMSO/H20 and assayed at 25 μM where the final DMSO concentration was 2%) was added to the testing wells. The ATP/33P-ATP solution (10 μL: containing 50 μM unlabeled ATP and 1 μCi 33P-ATP) was added to all wells and the completed plates were mixed and incubated at 30 °C for 30 minutes. Ice-cold 50% TCA/10 mM sodium phosphate (60 μL) was added to each well and the plates were kept on ice for 15 minutes.
The contents of each well were transferred to the wells of a 96-well filterplate (Millipore, MultiScreen- DP) and the filterplate was placed on a vacuum manifold, fitted with a waste collection tray. The wells were washed five times with 10% TCA/10 mM sodium phosphate (200 μL) under vacuum. MicroScint-20 scintillant was added, the plates were sealed using Topseal-S sheets and counted in a Packard TopCount scintillation counter using a 33P liquid program with color quench correction, where the output is in color quench-corrected cpm.
The % inhibition of the test compounds was calculated by the following formula: % inhibition = [1- (sample -BKG) / (CTRL-BKG) ] x 100. Compound 1 inhibited 44% of the activity of p38 at 20 μM.
2. In Vitro Cell Assay
In addition to the enzyme assay, many of the compounds of the invention were tested in an in vitro whole cell assay using peripheral blood mononuclear cells ("PBMC") which were obtained from human blood as follows. Freshly obtained venous blood was anticoagulated with heparin, diluted with an equal volume of phosphate buffered saline ("PBS") and placed in a sterile tube or other container. Aliquots (30 mL) of this mixture were transferred to centrifuge tubes which were underlaid with Ficoll-Hypaque (15 mL) . The prepared tubes were centrifuged at 400 x g without braking for 30 minutes at room temperature. Approximately 1/2 to 2/3 of the platelet layer above the mononuclear cell band was removed with a pipet. The majority of the mononuclear cell layer was carefully removed using a pipet and these PBMC s were diluted with PBS and spun at 600 x g for 15 minutes. The resulting PBMC s were washed with another portion of PBS and spun at 400 x g for 10 minutes at room temperature. The recovered pellets were diluted in low endotoxin RPMI / 1% FCS culture medium and gave a cell concentration of 0.5-2.0 x 10⁶ PBMC/ mL. A small volume of the suspension was removed for counting on a hemocytometer and the remaining preparation was centrifuged at 200 x g for 15 minutes at room temperature. The recovered pelleted PBM s were resuspended in RPMI / 1% FCS to a concentration of 1.67 x 10⁶/mL.
To run the assay, the PBMC suspension (180 μL) was transferred to duplicate wells of a 96-well flat-bottom microtiter plate and incubated for 1 hour at 37 °C. A solution of test compound (10 μL: prepared at 20 x the desired final concentration) was added to each well and the plate was incubated for 1 hour at 37 °C. A solution (10 μL) of LPS in RPMI / 1% FCS (200 ng/mL) was added and the wells were incubated overnight at 37°C. The supernatant (100 μL) was removed from each well and diluted with RPMI / 1% FCS (400 μL) . The samples were analyzed for TNF-α using a commercial ELISA kit (Genzyme) .
The anti IL-lβ activity of certain compounds of the invention was determined by the following in vitro assay. Plastic-adherent cells were prepared for PBMC. Briefly, PBMCs were added to the wells of a 96-well plate as above, incubated for lh at 37 °C, and the adherent cells prepared by gently re-suspending the non-adherent cells with a pipettor, removing and discarding the supernatant, and gently washing the wells 3 times with 200 μL of culture medium. Additional culture medium (180 μL) was added to the wells after the final wash. Compound addition, LPS stimulation, incubation and supernatant harvest were as for TNF-α. Supernatants were assayed for interleukin-lβ using a commercial ELISA (Genzyme) and IC5o values were determined. Compound 1 (described below) inhibited the production of IL-lβ at IC50 values of 400, 124, and 87 nM.
3. In Vivo Assay The ability of the compounds of Formula I to inhibit LPS-induced TNF-α production was demonstrated in the following in vivo rodent assays. Mice (BALB / cJ females, Jackson Laboratories) or rats (Lewis males, Charles River) were fasted for 30 minutes prior to oral dosing with 5-10 mL/kg of test compound at 5-50 mg/kg. Thirty minutes after dosing, the animals were injected intraperitoneally with LPS at 1 mg/kg and returned to their cages for 1 hour. Animals were anesthetized by C0 , exsanguinated by cardiac puncture and whole blood collected (0.1-0.7 mL) . The blood was allowed to clot and serum was transferred to a centrifuge tube. This sample was centrifuged, serum was collected, aliquoted and frozen at -80 °C. Samples were tested by commercial ELISA' s for TNF-α (Endogen for mouse TNF-α and Biosource for rat TNF-α) . In the mouse, compound 1 inhibited TNF-α 91% at 25 mg/kg and 74% at 10 mg/kg.
Certain compounds of the invention are listed in Tables A and B. The compounds were tested for their ability to inhibit TNF-α in vitro. The data are listed as an IC5o or as the % inhibition at a given concentration.
TABLE A
Figure imgf000024_0001
TNF-α
Figure imgf000024_0002
1 PhCH2 H # 7.0
2 Ph H # 20.0
3 H H # 60.0
4 CH3(CH2)3 H # 45.0
5 H Ph(CH2)3 # 1000
6 H (CH3)2N(CH2) 2# 2500
7 Ph(CH2)2 H # 50.0
8 H # P (CH2)3 200
# = absent
TABLE B
Figure imgf000025_0001
Cpd, Rα R2 R3 IC50 nm
9 H H # 15.0
10 PhCH2 H # >400
11 H Ph(CH2)3 # 0% @ 400 nM

Claims

What is claimed is:
1. A compound having the structure
Figure imgf000026_0001
or a pharmaceutically acceptable salt thereof, wherein:
(a) Ri, R2 and R3 are independently selected from the group consisting of (i) hydrogen, (ii) Cι-5alkyl, (iii) Cι_5alkylamino, (iv) diCι-5alkylamino, (v) a phenyl substituted with one or more of hydrogen, halogen, Ci-salkyl, and trihaloCι_5alkyl, and (vi) a phenylCι-5alkyl substituted with one or more of hydrogen, halogen, Cι_5alkyl, and trihaloCι_5alkyl;
(b) rings 1 and 2 are each independently substituted with one or more substituents selected from the group consisting of hydrogen, halogen, Ci-salkyl, and trihaloCι-5alkyl;
(c) A and B are independently nitrogen or carbon, at least one of A and B being nitrogen;
(d) D and E are nitrogen, with the proviso that (i) a double bond exists between the non-aryl carbon and either D or E, (ii) R2 is absent if the double bond exists between the non-aryl carbon and D, and (iii) R3 is absent if the double bond exists between the non-aryl carbon and E; and (e) the compound is neither 1, 6-dihydro-7- (4- pyridyl) -8- (4-fluorophenyl) -2-phenylmethyl- pyrrolo [3, 2-e]benzimidazole, nor 3, 6-dihydro-8- (4-fluorophenyl) -3- (3-phenylpropyl) -7- (4- pyridyl) -pyrrolo [3, 2-e]benzimidazole.
2. The compound of claim 1, wherein D and E are both nitrogen.
3. The compound of claim 1, wherein A is nitrogen and B is carbon.
4. The compound of claim 1, wherein D, E and A are nitrogen and B is carbon.
5. The compound of claim 1, wherein the compound is 1, 6-dihydro-7- (4-fluorophenyl) -8- (4-pyridyl) -2- phenyl-pyrrolo [3, 2-e]benzimidazole.
6. The compound of claim 1, wherein the compound is 1, 6-dihydro-7- (4-fluorophenyl) -8- (4-pyridyl) -2- butyl-pyrrolo [3, 2-e] benzimidazole.
7. The compound of claim 1, wherein the compound is 1, 6-dihydro-7- (4-fluorophenyl) -8- (4-pyridyl) -2- (2- phenylethyl) -pyrrolo [3, 2-e] benzimidazole.
8. The compound of claim 1, wherein the compound is 1, 6-dihydro-7- (4-pyridyl) -8- (4-fluorophenyl) - pyrrolo [3, 2-e] enzimidazole.
9. The compound of claim 1, wherein the compound is 1, 6-dihydro-7- (4-fluorophenyl) -8- (4-pyridyl) -2- phenylmethyl-pyrrolo [3, 2-e] benzimidazole.
10. A pharmaceutical composition comprising the compound of claim 1, and a pharmaceutically acceptable carrier.
11. A method of treating a subject having an inflammatory disease, which comprises administering to the subject a therapeutically effective dose of the pharmaceutical composition of claim 10.
12. The method of claim 11, wherein the disease is selected from the group consisting of rheumatoid arthritis, insulin-dependent diabetes mellitus, multiple sclerosis, myasthenia gravis, Crohn' s disease, autoimmune nephritis, primary biliary cirrhosis and psoriasis, acute pancreatitis, allograph rejection, allergic inflammation, inflammatory bowel disease, septic shock, osteoporosis, osteoarthritis, and cognitive deficits induced by neuronal inflammation.
13. The method of claim 12, wherein the disease is rheumatoid arthritis.
4. A method of making the compound of claim 1, which comprises the step of contacting a first compound having the structure
Figure imgf000029_0001
with a second compound having the structure
Figure imgf000029_0002
under conditions permitting a Fischer indolization between the first and second compounds.
PCT/US1999/004085 1998-02-26 1999-02-25 Substituted pyrrolobenzimidazoles for treating inflammatory diseases Ceased WO1999043680A1 (en)

Priority Applications (6)

Application Number Priority Date Filing Date Title
JP2000533434A JP4570777B2 (en) 1998-02-26 1999-02-25 Substituted pyrrolobenzimidazoles for treating inflammatory diseases
DE69917005T DE69917005T2 (en) 1998-02-26 1999-02-25 SUBSTITUTED PYRROLOBENZIMIDAZOLE DERIVATIVES FOR INFLAMMATION INHIBITION
EP99908456A EP1060181B1 (en) 1998-02-26 1999-02-25 Substituted pyrrolobenzimidazoles for treating inflammatory diseases
CA002321559A CA2321559A1 (en) 1998-02-26 1999-02-25 Substituted pyrrolobenzimidazoles for treating inflammatory diseases
AU27888/99A AU759297B2 (en) 1998-02-26 1999-02-25 Substituted pyrrolobenzimidazoles for treating inflammatory diseases
AT99908456T ATE266029T1 (en) 1998-02-26 1999-02-25 SUBSTITUTED PYRROLOBENZIMIDAZOLE DERIVATIVES FOR ANTI-INFLAMMATORY

Applications Claiming Priority (2)

Application Number Priority Date Filing Date Title
US7606398P 1998-02-26 1998-02-26
US60/076,063 1998-02-26

Publications (1)

Publication Number Publication Date
WO1999043680A1 true WO1999043680A1 (en) 1999-09-02

Family

ID=22129714

Family Applications (1)

Application Number Title Priority Date Filing Date
PCT/US1999/004085 Ceased WO1999043680A1 (en) 1998-02-26 1999-02-25 Substituted pyrrolobenzimidazoles for treating inflammatory diseases

Country Status (8)

Country Link
US (2) US6147096A (en)
EP (1) EP1060181B1 (en)
JP (1) JP4570777B2 (en)
AT (1) ATE266029T1 (en)
AU (1) AU759297B2 (en)
CA (1) CA2321559A1 (en)
DE (1) DE69917005T2 (en)
WO (1) WO1999043680A1 (en)

Cited By (6)

* Cited by examiner, † Cited by third party
Publication number Priority date Publication date Assignee Title
EP1707205A2 (en) 2002-07-09 2006-10-04 Boehringer Ingelheim Pharma GmbH & Co. KG Pharmaceutical compositions of anticholinergics and p38 kinase inhibitors in the treatment of respiratory diseases
WO2008142031A1 (en) 2007-05-18 2008-11-27 Institut Curie P38alpha as a therapeutic target in bladder carcinoma
EP2116245A2 (en) 2004-08-07 2009-11-11 Boehringer Ingelheim International GmbH EGFR kinase inhibitor combinations for treating respiratory and gastrointestinal disorders
US7737279B2 (en) 2005-05-10 2010-06-15 Bristol-Myers Squibb Company 1,6-dihydro-1,3,5,6-tetraaza-as-indacene based tricyclic compounds and pharmaceutical compositions comprising same
EP2384751A1 (en) 2004-12-24 2011-11-09 Boehringer Ingelheim International Gmbh Medicaments for the treatment or prevention of fibrotic diseases
WO2022022489A1 (en) * 2020-07-27 2022-02-03 江苏恒瑞医药股份有限公司 Indole fused ring derivative, preparation method therefor and use thereof in medicine

Families Citing this family (6)

* Cited by examiner, † Cited by third party
Publication number Priority date Publication date Assignee Title
DE10121215A1 (en) * 2001-04-30 2002-10-31 Merck Patent Gmbh Dihydro-imidazo [4,5-e] indole and 7H-pyrrolo [3,2-f] quinoxaline derivatives as nicotinic acetylcholine receptor ligands and / or serotonergic ligands
US6908899B2 (en) * 2001-08-17 2005-06-21 U.S. Dept. Of Veterans Affairs Pro-inflammatory fibrinopeptide
EP1631145A4 (en) * 2003-06-06 2009-12-23 Univ Pennsylvania P38 KINASEHEMMER COMPOSITION AND METHOD FOR THEIR USE
EP1676574A3 (en) 2004-12-30 2006-07-26 Johnson & Johnson Vision Care, Inc. Methods for promoting survival of transplanted tissues and cells
WO2015173788A1 (en) 2014-05-16 2015-11-19 Westfaelische Wilhelms-Universitaet Muenster Novel anti-infective strategy against influenza virus and s. aureus coinfections
WO2023280911A1 (en) 2021-07-06 2023-01-12 Westfälische Wilhelms-Universität Münster P38-inhibitors for the treatment of coronavirus infections and/or covid-19 cytokine storm

Citations (1)

* Cited by examiner, † Cited by third party
Publication number Priority date Publication date Assignee Title
WO1993014081A1 (en) * 1992-01-13 1993-07-22 Smithkline Beecham Corporation Imidazole derivatives and their use as cytokine inhibitors

Patent Citations (1)

* Cited by examiner, † Cited by third party
Publication number Priority date Publication date Assignee Title
WO1993014081A1 (en) * 1992-01-13 1993-07-22 Smithkline Beecham Corporation Imidazole derivatives and their use as cytokine inhibitors

Non-Patent Citations (2)

* Cited by examiner, † Cited by third party
Title
CHEMICAL ABSTRACTS, vol. 81, no. 5, 5 August 1974, Columbus, Ohio, US; abstract no. 25484, YOSHIDA ET AL: "Synthesis of antiinflammatory compounds. II. Synthesis of ring-fused pyrrole derivatives." page 417; column 1; XP002104371 *
YAKUGAKU ZASSHI, vol. 94, no. 2, 1974, pages 199 - 203 *

Cited By (8)

* Cited by examiner, † Cited by third party
Publication number Priority date Publication date Assignee Title
EP1707205A2 (en) 2002-07-09 2006-10-04 Boehringer Ingelheim Pharma GmbH & Co. KG Pharmaceutical compositions of anticholinergics and p38 kinase inhibitors in the treatment of respiratory diseases
EP2116245A2 (en) 2004-08-07 2009-11-11 Boehringer Ingelheim International GmbH EGFR kinase inhibitor combinations for treating respiratory and gastrointestinal disorders
EP2384751A1 (en) 2004-12-24 2011-11-09 Boehringer Ingelheim International Gmbh Medicaments for the treatment or prevention of fibrotic diseases
EP2878297A1 (en) 2004-12-24 2015-06-03 Boehringer Ingelheim International GmbH Medicaments for the treatment or prevention of fibrotic diseases
US7737279B2 (en) 2005-05-10 2010-06-15 Bristol-Myers Squibb Company 1,6-dihydro-1,3,5,6-tetraaza-as-indacene based tricyclic compounds and pharmaceutical compositions comprising same
US8268855B2 (en) 2005-05-10 2012-09-18 Bristol-Myers Squibb Company 1,6-dihydro-1,3,5,6-tetraaza-as-indacene based tricyclic compounds and pharmaceutical compositions comprising same
WO2008142031A1 (en) 2007-05-18 2008-11-27 Institut Curie P38alpha as a therapeutic target in bladder carcinoma
WO2022022489A1 (en) * 2020-07-27 2022-02-03 江苏恒瑞医药股份有限公司 Indole fused ring derivative, preparation method therefor and use thereof in medicine

Also Published As

Publication number Publication date
EP1060181A1 (en) 2000-12-20
AU2788899A (en) 1999-09-15
US6469174B1 (en) 2002-10-22
AU759297B2 (en) 2003-04-10
US6147096A (en) 2000-11-14
DE69917005T2 (en) 2005-03-24
EP1060181B1 (en) 2004-05-06
CA2321559A1 (en) 1999-09-02
JP2002504555A (en) 2002-02-12
DE69917005D1 (en) 2004-06-09
JP4570777B2 (en) 2010-10-27
ATE266029T1 (en) 2004-05-15

Similar Documents

Publication Publication Date Title
DE69816109T2 (en) SUBSTITUTED IMIDAZOLES FOR TREATING INFLAMMABLE DISEASES
US6610697B1 (en) Substituted 2-aryl-3-(heteroaryl)-imidazo[1,2-a]pyrimidines, and related pharmaceutical compositions and methods
WO1998047899A1 (en) Substituted pyrrolopyridines useful in the treatment of inflammatory diseases
WO1999003837A1 (en) 2-substituted imidazoles useful in the treatment of inflammatory diseases
EP1060181B1 (en) Substituted pyrrolobenzimidazoles for treating inflammatory diseases
EP1178981B1 (en) Substituted 3-pyridyl-4-arylpyrroles, and related therapeutic and prophylactic methods
US5817668A (en) S-heteroatom containing alkyl substituted-3-oxo-pyrido 1,2-A!benzimidozole-4-carboxamide derivatives useful in treating central nervous system disorders
CZ376999A3 (en) Substituted imidazoles useful in therapy of inflammatory diseases
AU2006202900A1 (en) Substituted imidazoles useful in the treatment of inflammatory diseases

Legal Events

Date Code Title Description
AK Designated states

Kind code of ref document: A1

Designated state(s): AL AM AT AU AZ BA BB BG BR BY CA CH CN CU CZ DE DK EE ES FI GB GE GH GM HR HU ID IL IS JP KE KG KP KR KZ LC LK LR LS LT LU LV MD MG MK MN MW MX NO NZ PL PT RO RU SD SE SG SI SK SL TJ TM TR TT UA UG UZ VN YU ZW

AL Designated countries for regional patents

Kind code of ref document: A1

Designated state(s): GH GM KE LS MW SD SL SZ UG ZW AM AZ BY KG KZ MD RU TJ TM AT BE CH CY DE DK ES FI FR GB GR IE IT LU MC NL PT SE BF BJ CF CG CI CM GA GN GW ML MR NE SN TD TG

121 Ep: the epo has been informed by wipo that ep was designated in this application
DFPE Request for preliminary examination filed prior to expiration of 19th month from priority date (pct application filed before 20040101)
WWE Wipo information: entry into national phase

Ref document number: 27888/99

Country of ref document: AU

ENP Entry into the national phase

Ref country code: JP

Ref document number: 2000 533434

Kind code of ref document: A

Format of ref document f/p: F

ENP Entry into the national phase

Ref document number: 2321559

Country of ref document: CA

Ref country code: CA

Ref document number: 2321559

Kind code of ref document: A

Format of ref document f/p: F

NENP Non-entry into the national phase

Ref country code: KR

WWE Wipo information: entry into national phase

Ref document number: 1999908456

Country of ref document: EP

WWP Wipo information: published in national office

Ref document number: 1999908456

Country of ref document: EP

REG Reference to national code

Ref country code: DE

Ref legal event code: 8642

WWG Wipo information: grant in national office

Ref document number: 27888/99

Country of ref document: AU

WWG Wipo information: grant in national office

Ref document number: 1999908456

Country of ref document: EP