WO2000023414A2 - Synthesis of optically active cyclohexylphenylglycolic acid and its esters - Google Patents
Synthesis of optically active cyclohexylphenylglycolic acid and its esters Download PDFInfo
- Publication number
- WO2000023414A2 WO2000023414A2 PCT/US1999/024721 US9924721W WO0023414A2 WO 2000023414 A2 WO2000023414 A2 WO 2000023414A2 US 9924721 W US9924721 W US 9924721W WO 0023414 A2 WO0023414 A2 WO 0023414A2
- Authority
- WO
- WIPO (PCT)
- Prior art keywords
- single enantiomer
- glycolic acid
- solution
- enantiomer
- cyclohexylphenyl glycolic
- Prior art date
- Legal status (The legal status is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the status listed.)
- Ceased
Links
- 0 *OC(C(c1ccccc1)(c1ccccc1)O)=O Chemical compound *OC(C(c1ccccc1)(c1ccccc1)O)=O 0.000 description 6
- YTRNSQPXEDGWMR-CQSZACIVSA-N O[C@](C1CCCCC1)(C(O)=O)c1ccccc1 Chemical compound O[C@](C1CCCCC1)(C(O)=O)c1ccccc1 YTRNSQPXEDGWMR-CQSZACIVSA-N 0.000 description 2
- DWKUEXWNTKVICP-DTQAZKPQSA-N OC(/C(/c1ccccc1)=C/CCC1CCCCC1)=O Chemical compound OC(/C(/c1ccccc1)=C/CCC1CCCCC1)=O DWKUEXWNTKVICP-DTQAZKPQSA-N 0.000 description 1
- YTRNSQPXEDGWMR-UHFFFAOYSA-N OC(C1CCCCC1)(C(O)=O)c1ccccc1 Chemical compound OC(C1CCCCC1)(C(O)=O)c1ccccc1 YTRNSQPXEDGWMR-UHFFFAOYSA-N 0.000 description 1
Classifications
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07C—ACYCLIC OR CARBOCYCLIC COMPOUNDS
- C07C69/00—Esters of carboxylic acids; Esters of carbonic or haloformic acids
- C07C69/96—Esters of carbonic or haloformic acids
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07C—ACYCLIC OR CARBOCYCLIC COMPOUNDS
- C07C213/00—Preparation of compounds containing amino and hydroxy, amino and etherified hydroxy or amino and esterified hydroxy groups bound to the same carbon skeleton
- C07C213/06—Preparation of compounds containing amino and hydroxy, amino and etherified hydroxy or amino and esterified hydroxy groups bound to the same carbon skeleton from hydroxy amines by reactions involving the etherification or esterification of hydroxy groups
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07C—ACYCLIC OR CARBOCYCLIC COMPOUNDS
- C07C219/00—Compounds containing amino and esterified hydroxy groups bound to the same carbon skeleton
- C07C219/02—Compounds containing amino and esterified hydroxy groups bound to the same carbon skeleton having esterified hydroxy groups and amino groups bound to acyclic carbon atoms of the same carbon skeleton
- C07C219/20—Compounds containing amino and esterified hydroxy groups bound to the same carbon skeleton having esterified hydroxy groups and amino groups bound to acyclic carbon atoms of the same carbon skeleton the carbon skeleton being unsaturated
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07C—ACYCLIC OR CARBOCYCLIC COMPOUNDS
- C07C229/00—Compounds containing amino and carboxyl groups bound to the same carbon skeleton
- C07C229/02—Compounds containing amino and carboxyl groups bound to the same carbon skeleton having amino and carboxyl groups bound to acyclic carbon atoms of the same carbon skeleton
- C07C229/34—Compounds containing amino and carboxyl groups bound to the same carbon skeleton having amino and carboxyl groups bound to acyclic carbon atoms of the same carbon skeleton the carbon skeleton containing six-membered aromatic rings
- C07C229/36—Compounds containing amino and carboxyl groups bound to the same carbon skeleton having amino and carboxyl groups bound to acyclic carbon atoms of the same carbon skeleton the carbon skeleton containing six-membered aromatic rings with at least one amino group and one carboxyl group bound to the same carbon atom of the carbon skeleton
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07C—ACYCLIC OR CARBOCYCLIC COMPOUNDS
- C07C51/00—Preparation of carboxylic acids or their salts, halides or anhydrides
- C07C51/42—Separation; Purification; Stabilisation; Use of additives
- C07C51/487—Separation; Purification; Stabilisation; Use of additives by treatment giving rise to chemical modification
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07C—ACYCLIC OR CARBOCYCLIC COMPOUNDS
- C07C67/00—Preparation of carboxylic acid esters
- C07C67/12—Preparation of carboxylic acid esters from asymmetrical anhydrides
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07C—ACYCLIC OR CARBOCYCLIC COMPOUNDS
- C07C68/00—Preparation of esters of carbonic or haloformic acids
- C07C68/02—Preparation of esters of carbonic or haloformic acids from phosgene or haloformates
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07B—GENERAL METHODS OF ORGANIC CHEMISTRY; APPARATUS THEREFOR
- C07B2200/00—Indexing scheme relating to specific properties of organic compounds
- C07B2200/07—Optical isomers
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07C—ACYCLIC OR CARBOCYCLIC COMPOUNDS
- C07C2601/00—Systems containing only non-condensed rings
- C07C2601/12—Systems containing only non-condensed rings with a six-membered ring
- C07C2601/14—The ring being saturated
Definitions
- the present invention relates to a method for resolving cyclohexylphenyl glycolic acid and the preparation of optically active phenylcyclohexylglycolate esters.
- Cyclohexylphenyl glycolic acid (also referred to herein as "CHPGA") is used as a starting material for manufacturing compounds that have important biological and therapeutic activities.
- Such compounds include, for example, oxphencyclimine, oxyphenonium bromide, oxypyrronium bromide, oxysonium iodide, oxybutynin (4- diethylamino-2-butynyl phenylcyclohexylglycolate) and its metabolites, such as desethyloxybutynin (4-ethylamino-2-butynyl phenylcyclohexylglycolate).
- oxphencyclimine oxyphenonium bromide
- oxypyrronium bromide oxysonium iodide
- oxybutynin (4- diethylamino-2-butynyl phenylcyclohexylglycolate)
- its metabolites such as des
- the (S)-enantiomers of oxybutynin and desethyloxybutynin have been shown to provide a superior therapy in treating urinary incontinence, as disclosed in U.S. Patent Nos. 5,532,278 and 5,677,346.
- the (R) enantiomer of oxybutynin has also been suggested to be a useful drug candidate. [Noronha-Blob et al., J. Pharmacol. Exp. Ther. 256, 562-567 (1991)].
- Racemic CHPGA is generally prepared by one of two methods: (1) selective hydrogenation of phenyl mandelic acid or of phenyl mandelate esters, as shown in Scheme 1; or (2) cyclohexyl magnesium halide addition to phenylglyoxylate as shown in Scheme 2.
- Scheme 1 selective hydrogenation of phenyl mandelic acid or of phenyl mandelate esters, as shown in Scheme 1; or (2) cyclohexyl magnesium halide addition to phenylglyoxylate as shown in Scheme 2.
- R is hydrogen or lower alkyl.
- a potential alternative to asymmetric synthesis is resolution of racemic CHPGA. This has been accomplished on an analytical scale using resolving agents such as ephedrine, quinine, and (+) and (-)-amphetamine.
- resolving agents such as ephedrine, quinine, and (+) and (-)-amphetamine.
- resolving agents are expensive, making known processes for resolution as impractical as known asymmetric syntheses.
- resolution processes using these agents provide poor stereoselectivity. The poor stereoselectivity necessitates multiple recrystallization steps to isolate the single CHPGA enantiomer, which adds to the production costs of chiral pharmaceuticals made from these precursors. Therefore, a need exists for a more efficient and economic method for preparing the single enantiomers of CHPGA on a commercial scale.
- a more efficient and economic method for producing chiral oxybutynin and its related compounds on an industrial scale is therefore desirable.
- a preparative scale method for producing the enantiomers of CHPGA for use as precursors, as well as a more convenient and inexpensive method for subsequent esterification would meet this need.
- Such methods should provide high purity compounds in high chemical yields with few processing steps, making them practical for use in the commercial production of optically active oxybutynin and related compounds.
- the process of the invention for preparing a single enantiomer of cyclohexylphenyl glycolic acid comprises: (a) providing a solution comprising racemic cyclohexylphenyl glycolic acid and a single enantiomer of tyro sine methyl ester;
- the invention relates to the diastereomeric salts, (S)- cyclohexylphenyl glycolic acid E-tyrosine methyl ester salt and (R)-cyclohexylphenyl glycolic acid D-tyrosine methyl ester salt, which crystallize when L-tyrosine methyl ester or .D-tyrosine methyl ester, respectively, is combined with racemic cyclohexylphenyl glycolic acid in the above process.
- the invention relates to a process for preparing a single enantiomer of structure (II)
- R 1 is chosen from the group of hydrogen or -CH 2 R 2 , wherein R 2 is chosen from the group of azide, hydroxy, halo, or -NR 3 R 4 , wherein R 3 and R 4 are each independently lower alkyl, benzyl or methoxybenzyl .
- the synthetic method directly couples an enantiomer of CHPGA with a propargyl alcohol derivative to produce a single enantiomer of a propargyl ester of cylclohexylphenylglycolate.
- the method comprises:
- R 1 is as previously defined to produce the single enantiomer of structure (II).
- R 3 and R 4 be converted to hydrogen
- the product of the condensation of CHPGA with the butynol in which one or both of R 3 and R 4 is benzyl or methoxybenzyl may be deprotected by treatment with ⁇ -chloroethyl carbonochloridate.
- the invention relates to the preparation of the hydrochloride salt of a single enantiomer of structure (II) above, in which R 1 is -CH 2 R 2a and R 2a is -NR 3 R 4 .
- the process comprises preparing the single enantiomer of structure (II) according to the method set forth above, then separating the organic solvent containing the single enantiomer of structure (II) from the reaction mixture.
- the organic solvent is then exchanged with ethyl acetate to produce an ethyl acetate solution containing the single enantiomer of structure (II).
- the ethyl acetate solution is then concentrated and dried to contain about 20-25% by weight single enantiomer of structure (II) and ⁇ 0.3% by weight water.
- Methyl t-butyl ether is then added to provide an ethyl acetate/t-butyl ether solution, wherein the concentration of the single enantiomer of structure (II) is reduced by about one third.
- a mixture of HC1 in ethanol is added to the ethyl acetate/t-butyl ether solution to form a hydrochloride salt of the single enantiomer of structure (II).
- the invention relates to the preparation of a single enantiomer of structure (II) using the aforementioned coupling process in which the single enantiomer of cyclohexylphenyl glycolic acid of structure , (I) is obtained using the resolution process outlined above.
- R 5 is alkyl or alkenyl of one to ten carbons.
- R 5 is isobutyl. Both intermediates are useful in the preparation of optically pure isomers of CHPGA esters, such as oxybutynin.
- Advantages of the present resolution process over the prior art include: higher efficiency, fewer processing steps, and a more readily available resolving agent.
- Previous resolution processes utilizing ephedrine as the resolving agent require a multi-step crystallization process to obtain (S)-CHPGA (e.g., 98- 99% ee). The overall yield of these steps is approximately 30%.
- the process described herein using an enantiomer of tyrosine methyl ester gives a 42% yield of 99% ee diastereomer in a single resolution step.
- the advantages of the present synthetic method for producing oxybutynin and its analogs over the prior art methods include: a one step process from chiral CHPGA without isolation of any intermediates, and an improvement in yield and quality over the transesterification process shown in Scheme 3.
- the process can also be used to produce a variety of analogs not available through transesterification or alkoxy radical couplings.
- enantiomeric excess is related to the older term “optical purity” in that both are measures of the same phenomenon.
- the value of ee will be a number from 0 to 100, zero being racemic and 100 being pure, single enantiomer.
- a compound which in the past might have been called 98% optically pure is now more precisely described as 96% ee.; in other words, a 90% e.e. reflects the presence of 95% of one enantiomer and 5% of the other in the material in question.
- Alkyl refers to saturated hydrocarbon residues containing twenty or fewer carbons in straight or branched chains, as well as cyclic structures.
- Lower alkyl is 6 or fewer carbons.
- Alkenyl refers to hydrocarbon residues containing twenty or fewer carbons in straight or branched chains, as well as cyclic structures having at least one degree of unsaturation.
- Lower alkenyl is 6 or fewer carbons.
- the term “% by weight” is interchangeable with and has the same meaning as “wt %”.
- equiv.” refers to molar equivalents.
- the reactants and reagents used in the processes of the present invention described herein are readily available materials. Such materials can be conveniently prepared with conventional preparatory procedures or obtained from commercial sources.
- the present process for preparing a single enantiomer of cyclohexylphenyl glycolic acid comprises:
- the solution of step (a) is accomplished by dissolving a mixture of racemic cyclohexylphenyl glycolic acid and a single enantiomer of tyrosine methyl ester in acetonitrile and water by heating to form a solution and step (b) is accomplished by allowing the solution to cool, whereby a salt of primarily one diastereomer crystallizes.
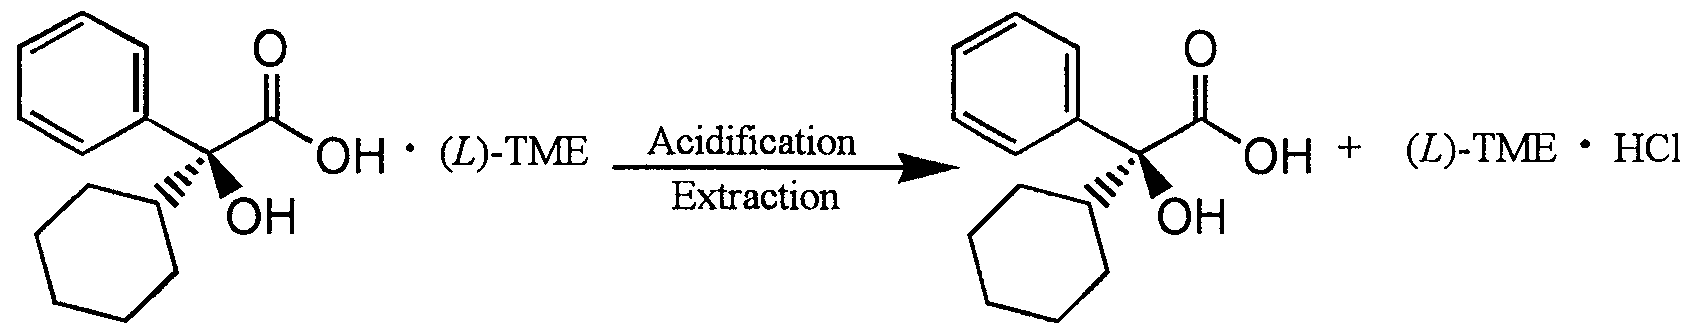
- Liberation of the single enantiomer of cyclohexylphenyl glycolic acid from the diastereomeric salt may be accomplished, by adding the diastereomeric salt to a mixture of toluene and aqueous mineral acid.
- the resulting toluene solution may then be reduced in volume and cooled.
- the single enantiomer of cyclohexylphenyl glycolic acid product may then be filtered off.
- tyrosine methyl ester may be recovered.
- the aspect of the invention relating to a synthetic method for directly coupling an enantiomer of CHPGA with a propargyl alcohol derivative to produce a single enantiomer of a propargyl ester of cylclohexylphenylglycolate includes a step in which the carboxy group of a single enantiomer of cyclohexylphenyl glycolic acid is activated to form an activated carboxy compound.
- Methods for activating carboxyl toward displacement by alcohols are known in the art for preparing esters.
- the starting material in this case contains an unprotected alcohol function, it is surprising that the activation and condensation can be carried out with sufficient specificity to provide a product which is not an intractable mixture of self- condensation products of the glycolic acid component.
- the single enantiomer of cyclohexylphenyl glycolic acid is reacted with an alkyl chloroformate in an organic solvent to form a mixed anhydride; and a sidechain propargyl alcohol derivative of structure (III) is added to the mixed anhydride.
- the alkyl chloroformate is isobutylchloroformate.
- the resolution process of the present invention provides an inexpensive and efficient method for preparing a single enantiomer from racemic CHPGA via the formation of the diastereomeric salt with (L) or (D) -tyrosine methyl ester, also referred to herein as "(Z) or (D)-TME".
- the process consists of three parts, which are depicted and described below: Part 1: Preparation of (S)-CHPGA-(Z)-TME diastereomeric salt or (R)-CHPGA-(D)-TME diastereomeric salt; Part 2:
- the racemic starting material, (S, R)- cyclohexylphenyl glycolic acid can be prepared by the process described above, i.e. (1) selective hydrogenation of phenyl mandelic acid or of phenyl mandelate esters or (2) cyclohexyl magnesium halide addition to phenylglyoxylate.
- Mandelic acid and phenylglyoxylic acid, also known as benzoylformic acid are commercially available.
- Phenyl mandelic acid may be prepared by Grignard addition of phenyl magnesium bromide to diethyl oxalate followed by hydrolysis.
- the (L) enantiomer of tyrosine methyl ester is also readily available from commercial sources, as is (Z ) )-tyrosine, which can then be esterified to produce (_9)-tyrosine methyl ester using conventional techniques, such as acid-catalyzed esterification with methanol.
- the diastereomer of the present process is produced by dissolving racemic
- a suitable solvent for example, racemic CHPGA and about 0.5 molar equivalents of (Z)-tyrosine methyl ester or (Z))-tyrosine methyl ester can be dissolved in a mixture of acetonitrile and water.
- solvent is about 10 wt % water in acetonitrile, solution may be achieved by heating, preferably by heating to reflux (approximately 78° C).
- the CHPGA enantiomer produced, (S) or (R)-CHPGA is liberated from the diastereomeric salt.
- the (S)-CHPGA-(E)- TME salt from Part 1 is added to and dissolved to form a solution which is about 15 wt % substrate in toluene.
- the solution is treated with an excess of dilute mineral acid, such as 1.1 equivalents of 0.5 M HC1 or H 2 SO 4 .
- diastereomeric salt essentially all the TME enantiomer is converted to the hydrochloride salt.
- the diastereomeric salt mixture may be heated to about 40-50° C for about 10 minutes to facilitate dissolution of the solids.
- a phase split yields an aqueous solution containing (Z)-TME-HCl and an organic solution of (S)-CHPGA in toluene.
- the aqueous phase is separated from the organic solution and saved for recovery of the tryrosine methyl ester in Step 3 below.
- a common method of separation which may be used throughout the processes described herein, is gravitational settling followed by drainage of the aqueous phase through a tap in the bottom of the reaction vessel.
- the toluene organic phase containing (S)-CHPGA may be washed a second time with mineral acid, as specified above, and heated.
- the organic phase and aqueous phase are then separated, and the aqueous phase is discarded along with the rag layer, i.e. the layer separating the two phases.
- the retained toluene organic phase is then preferably concentrated, typically by vacuum distillation, to a weight that is about 2.1 to 2.3 times the weight of the diastereomeric salt originally present, followed by gradual cooling to 0-5° C to initiate crystallization of the single (S) enantiomer of CHPGA, as indicated by the formation of a thick slurry.
- the slurry is cooled for at least an hour to ensure that crystallization is complete, then filtered to isolate (S)-CHPGA.
- the (S)-CHPGA cake is then dried under vacuum while heating to a temperature of about (40-45° C).
- Part 2 is cooled, preferably to about 0-5° C. While maintaining the cooling temperature, the aqueous solution is titrated with 0.5M NaOH to a pH of approximately 9.0. Typically, a thin slurry will form as the TME enantiomer precipitates.
- the TME enantiomer is isolated by filtration, washing with deionized water, and drying under vacuum at a temperature of about (40-50° C).
- Part 1 447.0 g toluene, 339.2 g 0.5M HC1 (1.1 equiv.) and heated to 40-50°C while stirring until the solids dissolved (about 10 min). While maintaining the temperature at 40-50° C, the organic and aqueous phases separated after about 10 minutes. The phases were divided, and the aqueous (bottom) phase containing (L)- TME-HC1 was saved for recovery in Part 3 below. Approximately 370 g aqueous phase was recovered.
- the slurry was filtered to recover the (S)-CHPGA.
- the synthesis of a single enantiomer of oxybutynin and oxybutynin analogs comprises coupling an enantiomer of cyclohexylphenyl glycolic acid with a propargyl alcohol derivative utilizing carboxylic acid activation.
- Optically active CHPGA may be prepared either by the resolution process described above or by asymmetric methods.
- the present invention also provides a process for converting the aforementioned enantiomers of oxybutynin and oxybutynin analogs to their corresponding hydrochloride salts.
- the synthetic process consists of two reactions, which are depicted and described below: Part 1: Formation of the Mixed Anhydride; Part 2: Formation of (S) or (R) oxybutynin and its related compounds. Again for ease in understanding, the (S) enantiomeric series is depicted, although the (R) series is produced similarly.
- the reaction is preferably carried out in an inert atmosphere, such as nitrogen or argon, and the reaction solution is stirred using conventional techniques.
- IBCF isobutyl chloroformate
- isopropenylchloroformate and 2-ethylhexylchloroformate may instead be used.
- the amount of alkyl chloroformate used in the reaction is preferably about 1.2 equivalents with respect to the CHPGA enantiomer.
- the reaction proceeds in the presence of a tertiary amine (2.5 equiv.), such as triethylamine (TEA), 4-N,N-dimethylaminopyridine (DMAP), pyridine, diisopropylethylamine, diethylmethylamine, ⁇ -methylpiperidine or ⁇ - methylmorpholine, which scavenges the HC1 produced.
- a tertiary amine such as triethylamine (TEA), 4-N,N-dimethylaminopyridine (DMAP), pyridine, diisopropylethylamine, diethylmethylamine, ⁇ -methylpiperidine or ⁇ - methylmorpholine, which scavenges the HC1 produced.
- Organic solvents that may be used include, but are not limited to cyclohexane, heptane, toluene, tetrahydrofuran (THF), ethylene glycol dimethyl ether (DME
- a sidechain propargyl alcohol derivative of formula (III), wherein R 1 is as previously defined, is added to the mixed anhydride contained in the reaction mixture to produce the single enantiomer of oxybutynin or analog thereof (II).
- About 1.3 equivalents of the formula (III) compound relative to (S) or (R)-CHPGA is sufficient.
- the reaction mixture is heated to reflux at a temperature of about 65-80° C, but more preferably about 70-75° C, until the reaction is complete, as determined by HPLC.
- the propargyl alcohol derivative of formula (III) is a 4- amino propargyl alcohol derivative, wherein R 1 is represented as -CH 2 R 2 ; R 2 is - NR 3 R 4 ; and R 3 and R 4 are each independently lower alkyl, benzyl or methoxybenzyl.
- the compound of formula (III) is most preferably 4-N,N-diethylamino butynol (4-N,N-DEB), where R 3 and R 4 are each ethyl. Reaction of the mixed anhydride with 4-N.N-DEB produces the single enantiomer of oxybutynin, i.e.
- (S) or (R)-4-diethylamino-2-butynyl phenylcyclohexylglycolate Another preferred embodiment is the reaction using an ⁇ -protected 4-N-ethylamino butynol, such as ⁇ -ethyl- ⁇ -(4-methoxybenzyl)butynol, as the propargyl alcohol derivative and then cleaving the protecting group (by methods well known in the art) to produce (S) or (R)-4-ethylamino-2-butynyl phenylcyclohexylglycolate, also known as desethyloxybutynin.
- an ⁇ -protected 4-N-ethylamino butynol such as ⁇ -ethyl- ⁇ -(4-methoxybenzyl)butynol
- R 3 is ethyl
- R 4 is converted to hydrogen in formula (III).
- Suitable protecting groups are described in Greene and Wuts Protecting Groups in Organic Synthesis. Second Edition Wiley, New York 1991, p. 362-371, which is incorporated herein by reference.
- the propargyl alcohol derivative of formula (III) is 4-N,N- ethylmethylamino butynol, which results in the formation of (Sf) or (R)-4- ethylmethylamino-2-butynyl phenylcyclohexylglycolate.
- R 3 is ethyl
- R 4 is methyl.
- R 1 is -CH 2 R 2
- R 2 is azide, hydroxy, or halo
- propargyl alcohol itself also known as 2-propyn-l-ol
- R 1 is hydrogen in formula (III).
- 4-N,N-Diethylamino butynol for use as the sidechain propargyl alcohol in the present invention may be prepared by reacting propargyl alcohol, paraformaldehyde, and diethylamine under standard Mannich conditions.
- Other amino and alkyl amino propargyl alcohol derivatives of structure (III) can be formed by the process disclosed in U.S. Patent No. 5,677,346.
- a secondary amine in which one or more substituents may be a protecting group, such as N-ethyl-4- methoxybenzenemethanamine for example, is reacted with propargyl alcohol and paraformaldehyde in the presence of cuprous chloride.
- the addition of ⁇ -chloroethyl carbonochloridate removes the protecting group.
- the 4-N-ethylaminobutynyl ester is the ultimate product.
- the remaining propargyl alcohol derivatives for use in the present invention are commercially available or can be synthesized by methods known in the art.
- the progress of the condensation of the mixed anhydride with the propargyl alcohol may be conveniently monitored by periodic HPLC analyses of the reaction mixture until the desired extent of conversion is reached.
- the reaction is preferably quenched by washing with 10-12% aqueous monobasic sodium phosphate and water. About 8.5 g of the phosphate per gram of enantiomeric CHPGA used is typical.
- the aqueous washes are then discarded.
- a final wash using deionized water may then be performed, after which the bottom aqueous phase is discarded.
- the retained organic phase containing the enantiomer of structure (II) in solution with the organic solvent can then be concentrated to remove most of the solvent, typically by vacuum distillation.
- the organic solvent containing the enantiomer of oxybutynin or one of its analogs (II) produced by the process outlined above (also referred to herein as "free base enantiomer (II)" is exchanged with ethyl acetate (EtOAc).
- EtOAc ethyl acetate
- the organic solvent is removed by vacuum distillation to contain about 20-25 wt % (S) or (R) enantiomer of structure (II), which is based on the theoretical amount of free base (II) formed from the coupling process above.
- Ethyl acetate is then added to obtain the original solution volume or weight. This step may be repeated substituting the removal of EtOAc for the organic solvent.
- the EtOAc solution may be filtered through a filtering agent, such as diatomaceous earth. The filter cake is washed with EtOAc as needed.
- the filtrate is then concentrated by vacuum distillation, for example, to contain about 20-25 wt % (theoretical) of free base enantiomer (II) and ⁇ 0.3 wt % water. To maximize product yield and purity and to encourage crystallization, most of the water should be removed from the solution. If the foregoing concentration processes are insufficient to reduce the water to ⁇ 0.3%, the vacuum distillation may be repeated with fresh solvent or a drying agent, such as magnesium sulfate may be employed. Water content can be determined by KF (Karl Fisher method).
- Methyl t-butyl ethyl (MTBE) is then added to the concentrated EtOAc solution to a volume that reduces the concentration by weight of the free base enantiomer (II) by about one third, or optimally to between about 6.5 and 8.5 wt %.
- the hydrochloride salt is then formed by the addition of HC1, while stirring. A slight excess of HC1 in ethanol, for example about 1.1 equivalents of 35-40 wt % HC1, is generally sufficient.
- the temperature may be increased to 35-45° C.
- the solution may be seeded with the hydrochloride salt of the enantiomer of structure (II).
- a slurry forms. If the slurry is cooled to about 0-5° C and this temperature maintained for about two hours, filtration provides a very good recovery of the hydrochloride salt of the enantiomer of structure (II).
- the filter cake is typically a white to off-white crystalline solid, which can then be washed with ambient temperature methyl t-butyl ether (at least 2.2 g MTBE per gram free base enantiomer (II)), followed by vacuum drying at 40- 50° C.
- Cyclohexane was removed from the organic phase by vacuum distillation to about 350 g ( ⁇ 22 wt % (S)-oxybutynin based on the theoretical amount (76.29 g) of (S)-oxybutynin free base formed).
- Ethyl acetate (EtOAc) was added to obtain the original solution volume of about 1000 mL (or about 83 Og), followed by vacuum distillation to 20-25 wt % (S)-oxybutynin.
- EtOAc was then added a second time to a volume of about 1000 mL (or about 830g). The batch was then polish filtered through about 5.0 g CELITE® while washing the filter cake with EtOAc as needed.
- the filtered mixture was concentrated and dried by vacuum distillation to 339 g ( ⁇ 22.5 wt % (S)-oxybutynin) and ⁇ 03 wt % water, as measured by KF. Based on the theoretical amount of (S)-oxybutynin free base (76.29 g), methyl t-butyl ether was added to adjust the (S)-oxybutynin free base concentration to 8.0 wt % (953 g). With agitation, 23 g of 37 wt % HC1 in EtOH (1.1 equiv.) were slowly added to the solution, while maintaining the temperature between 20 and 45° C.
- the temperature of the solution was then adjusted to 35-45° C, and the solution was seeded with about 500 mg (S)-oxybutynin-HCl crystals (approximately 10 mg of seeds per g (S)-CHPGA ). The temperature was maintained, and the solution was stirred for about one hour. A slurry formed, which was then cooled to 0-5 °C over a minimum of 1 hour and held for 2 hours. The slurry was then filtered to recover the (S)-oxybutynin-HCl. The filter cake was a white to off-white crystalline solid.
- Example 3 Isolation of the two carbonate intermediates A and B
- CHPGA cyclohexylphenylglycolic acid
- cyclohexane 100 mL
- triethylamine 7.4 mL, 0.053 mol
- isobutylchloroformate 5.5mL, 0.0426mol
- the slurry was allowed to stir at ambient temperature for approximately 0.5 h, at which time the reaction was quenched with a 10% aq. NaH 2 PO 4 (50 ml).
- the organic phase was separated from the aqueous phase and washed with 10% aq.
Landscapes
- Chemical & Material Sciences (AREA)
- Organic Chemistry (AREA)
- Chemical Kinetics & Catalysis (AREA)
- General Chemical & Material Sciences (AREA)
- Engineering & Computer Science (AREA)
- Oil, Petroleum & Natural Gas (AREA)
- Organic Low-Molecular-Weight Compounds And Preparation Thereof (AREA)
- Low-Molecular Organic Synthesis Reactions Using Catalysts (AREA)
Abstract
Description




Claims


Priority Applications (7)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| JP2000577142A JP2002527500A (en) | 1998-10-22 | 1999-10-22 | Optically active cyclohexylphenyl glycolic acid and its esters |
| DE69916134T DE69916134T2 (en) | 1998-10-22 | 1999-10-22 | PREPARATION OF OPTICALLY ACTIVE CYCLOHEXYLPHENYL GLYCOLIC ACID AND ITS ESTERS |
| EP99953250A EP1185498B1 (en) | 1998-10-22 | 1999-10-22 | Synthesis of optically active cyclohexylphenylglycolic acid and its esters |
| AU65222/99A AU766066B2 (en) | 1998-10-22 | 1999-10-22 | Synthesis of optically active cyclohexylphenylglycolic acid and its esters |
| AT99953250T ATE263134T1 (en) | 1998-10-22 | 1999-10-22 | PRODUCTION OF OPTICALLY ACTIVE CYCLOHEXYLPHENYL GLYCOLIC ACID AND ITS ESTERS |
| CA002347736A CA2347736A1 (en) | 1998-10-22 | 1999-10-22 | Synthesis of optically active cyclohexylphenylglycolic acid and its esters |
| AU2003271292A AU2003271292A1 (en) | 1998-10-22 | 2003-12-22 | Synthesis of optically active cyclohexylphenylglycolic acid and its esters |
Applications Claiming Priority (4)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| US09/177,456 US5973182A (en) | 1998-10-22 | 1998-10-22 | Carbonate Intermediates useful in the preparation of optically active cyclohexylphenylglycolate esters |
| US09/177,456 | 1998-10-22 | ||
| US09/211,646 US6140529A (en) | 1998-10-22 | 1998-12-15 | Synthesis of optically active cyclohexylphenylglycolate esters |
| US09/211,646 | 1998-12-15 |
Publications (2)
| Publication Number | Publication Date |
|---|---|
| WO2000023414A2 true WO2000023414A2 (en) | 2000-04-27 |
| WO2000023414A3 WO2000023414A3 (en) | 2001-12-13 |
Family
ID=26873314
Family Applications (1)
| Application Number | Title | Priority Date | Filing Date |
|---|---|---|---|
| PCT/US1999/024721 Ceased WO2000023414A2 (en) | 1998-10-22 | 1999-10-22 | Synthesis of optically active cyclohexylphenylglycolic acid and its esters |
Country Status (11)
| Country | Link |
|---|---|
| US (1) | US6140529A (en) |
| EP (3) | EP1185498B1 (en) |
| JP (1) | JP2002527500A (en) |
| AT (2) | ATE263134T1 (en) |
| AU (2) | AU766066B2 (en) |
| CA (1) | CA2347736A1 (en) |
| DE (2) | DE69932269T2 (en) |
| DK (2) | DK1424321T3 (en) |
| ES (2) | ES2215406T3 (en) |
| PT (2) | PT1424321E (en) |
| WO (1) | WO2000023414A2 (en) |
Cited By (2)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| US6777576B2 (en) | 2000-11-07 | 2004-08-17 | Sumika Fine Chemicals Co., Ltd. | Production method of 2-cyclohexyl-2-hydroxy-2-phenylacetic acid intermediate therefor and production method thereof |
| CN103012231A (en) * | 2012-12-14 | 2013-04-03 | 沈阳药科大学 | Preparation method and application of glycopyrronium bromide chiral antipode |
Families Citing this family (4)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| US6180823B1 (en) * | 1998-11-06 | 2001-01-30 | Sepracor Inc. | Stereoselective process for alkyl phenylglycolic acids |
| US6825378B2 (en) * | 2003-03-28 | 2004-11-30 | Council Of Scientific And Industrial Research | Process for the synthesis of enantiomerically pure cyclohexylphenyl glycolic acid |
| CN103772188B (en) * | 2014-02-25 | 2015-09-23 | 成都百事兴科技实业有限公司 | The preparation method of R-(+)-α-Cyclo hexyl mandelic acid |
| CR20220617A (en) | 2020-05-05 | 2023-02-09 | Apnimed Inc | Polymorphic forms of (r)-oxybutynin hydrochloride |
Family Cites Families (7)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| GB708370A (en) * | 1950-12-27 | 1954-05-05 | Albert Boehringer | Process for the production of scopolamine-bromobutylate |
| GB750156A (en) * | 1953-03-23 | 1956-06-13 | Wellcome Found | Improvements in cyclohexyl-phenyl-amino-propanols |
| US2837525A (en) * | 1955-07-12 | 1958-06-03 | Sterling Drug Inc | 1-thienyl-1-cycloalkyl (or aryl)-3-(aliphatic-tertiary-amino)-1-hydroxy-lower-alkanes and preparation thereof |
| NL267508A (en) * | 1960-07-26 | |||
| JP2651411B2 (en) * | 1988-06-13 | 1997-09-10 | 日本ヘキスト・マリオン・ルセル株式会社 | Preparation of alkylaminoacetylene glycolate |
| IL91377A (en) * | 1988-09-14 | 1996-09-12 | Nippon Shinyaku Co Ltd | Butynylamine glycolate derivatives |
| US6207852B1 (en) * | 1996-07-01 | 2001-03-27 | Bridge Pharma, Inc. | Smooth muscle spasmolytic agents, compositions and methods of use thereof |
-
1998
- 1998-12-15 US US09/211,646 patent/US6140529A/en not_active Expired - Lifetime
-
1999
- 1999-10-22 WO PCT/US1999/024721 patent/WO2000023414A2/en not_active Ceased
- 1999-10-22 PT PT03104950T patent/PT1424321E/en unknown
- 1999-10-22 ES ES99953250T patent/ES2215406T3/en not_active Expired - Lifetime
- 1999-10-22 CA CA002347736A patent/CA2347736A1/en not_active Abandoned
- 1999-10-22 PT PT99953250T patent/PT1185498E/en unknown
- 1999-10-22 JP JP2000577142A patent/JP2002527500A/en not_active Withdrawn
- 1999-10-22 DE DE69932269T patent/DE69932269T2/en not_active Expired - Fee Related
- 1999-10-22 DK DK03104950T patent/DK1424321T3/en active
- 1999-10-22 ES ES03104950T patent/ES2266726T3/en not_active Expired - Lifetime
- 1999-10-22 AT AT99953250T patent/ATE263134T1/en not_active IP Right Cessation
- 1999-10-22 AT AT03104950T patent/ATE332290T1/en not_active IP Right Cessation
- 1999-10-22 EP EP99953250A patent/EP1185498B1/en not_active Expired - Lifetime
- 1999-10-22 AU AU65222/99A patent/AU766066B2/en not_active Ceased
- 1999-10-22 EP EP03104950A patent/EP1424321B1/en not_active Expired - Lifetime
- 1999-10-22 DK DK99953250T patent/DK1185498T3/en active
- 1999-10-22 EP EP05301107A patent/EP1642883A3/en not_active Withdrawn
- 1999-10-22 DE DE69916134T patent/DE69916134T2/en not_active Expired - Fee Related
-
2003
- 2003-12-22 AU AU2003271292A patent/AU2003271292A1/en not_active Abandoned
Cited By (3)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| US6777576B2 (en) | 2000-11-07 | 2004-08-17 | Sumika Fine Chemicals Co., Ltd. | Production method of 2-cyclohexyl-2-hydroxy-2-phenylacetic acid intermediate therefor and production method thereof |
| CN103012231A (en) * | 2012-12-14 | 2013-04-03 | 沈阳药科大学 | Preparation method and application of glycopyrronium bromide chiral antipode |
| CN103012231B (en) * | 2012-12-14 | 2015-08-05 | 沈阳药科大学 | The preparation method and application of Glycopyrronium Bromide chiral enantiomer |
Also Published As
| Publication number | Publication date |
|---|---|
| EP1185498B1 (en) | 2004-03-31 |
| DE69932269D1 (en) | 2006-08-17 |
| ES2266726T3 (en) | 2007-03-01 |
| DE69916134D1 (en) | 2004-05-06 |
| DE69932269T2 (en) | 2006-11-23 |
| US6140529A (en) | 2000-10-31 |
| EP1185498A2 (en) | 2002-03-13 |
| DK1424321T3 (en) | 2006-07-31 |
| EP1424321A1 (en) | 2004-06-02 |
| AU766066B2 (en) | 2003-10-09 |
| PT1424321E (en) | 2006-09-29 |
| AU6522299A (en) | 2000-05-08 |
| PT1185498E (en) | 2004-06-30 |
| DE69916134T2 (en) | 2005-02-03 |
| EP1424321B1 (en) | 2006-07-05 |
| ATE263134T1 (en) | 2004-04-15 |
| ATE332290T1 (en) | 2006-07-15 |
| DK1185498T3 (en) | 2004-07-26 |
| AU2003271292A1 (en) | 2004-01-29 |
| ES2215406T3 (en) | 2004-10-01 |
| EP1642883A2 (en) | 2006-04-05 |
| JP2002527500A (en) | 2002-08-27 |
| CA2347736A1 (en) | 2000-04-27 |
| WO2000023414A3 (en) | 2001-12-13 |
| EP1642883A3 (en) | 2006-05-03 |
| AU2003271292A2 (en) | 2004-01-29 |
Similar Documents
| Publication | Publication Date | Title |
|---|---|---|
| JP3965440B2 (en) | Method for producing (S) -3- (aminomethyl) -5-methylhexanoic acid | |
| EP0830338B1 (en) | Method of making 3-(aminomethyl)-5-methylhexanoic acid | |
| US6090971A (en) | Resolution process for cyclohexylphenyl glycolic acid | |
| EP0435068B1 (en) | Process for producing optically active atenolol and intermediate thereof | |
| US6140529A (en) | Synthesis of optically active cyclohexylphenylglycolate esters | |
| US4571424A (en) | Optical resolution of racemic femoxetine | |
| AU2007267371B2 (en) | New synthesis of substituted hydroxymethyl phenols | |
| JPH06184069A (en) | Production of alpha-hydroxy-beta-aminocarboxylic acid | |
| JP2004534040A (en) | Stereoselective synthesis of 2-hydroxy-4-phenylbutyrate | |
| EP2448916B1 (en) | Production of trans-4-aminocyclopent-2-ene-1-carboxylic acid derivatives | |
| WO2007026373A2 (en) | Process for preparing rivastigmine | |
| EP0319024B1 (en) | Process for producing optically active alpha-isopropyl-p-chlorophenylacetic acid | |
| US7122696B2 (en) | Processes for preparation of N-protected-β-amino alcohols and N-protected-β-amino epoxides | |
| WO2005080329A2 (en) | Process for manufacture of racemic carvedilol | |
| EP0339618B1 (en) | Method for preparing optically active 3,4-dihydroxy butyric acid derivatives | |
| US20040039206A1 (en) | Process for resolving racemic mixtures of piperidine derivatives | |
| US20050065366A1 (en) | Method for producing optically active beta-phenylalanine compounds | |
| WO1992015575A1 (en) | Process and intermediate for certain bis-aza-bicyclic anxiolytic agents | |
| EP1862449A1 (en) | A shortened synthesis of substituted hydroxymethyl phenols |
Legal Events
| Date | Code | Title | Description |
|---|---|---|---|
| ENP | Entry into the national phase |
Ref country code: AU Ref document number: 1999 65222 Kind code of ref document: A Format of ref document f/p: F |
|
| AK | Designated states |
Kind code of ref document: A2 Designated state(s): AE AL AM AT AU AZ BA BB BG BR BY CA CH CN CR CU CZ DE DK DM EE ES FI GB GD GE GH GM HR HU ID IL IN IS JP KE KG KP KR KZ LC LK LR LS LT LU LV MD MG MK MN MW MX NO NZ PL PT RO RU SD SE SG SI SK SL TJ TM TR TT TZ UA UG UZ VN YU ZA ZW |
|
| AL | Designated countries for regional patents |
Kind code of ref document: A2 Designated state(s): GH GM KE LS MW SD SL SZ TZ UG ZW AM AZ BY KG KZ MD RU TJ TM AT BE CH CY DE DK ES FI FR GB GR IE IT LU MC NL PT SE BF BJ CF CG CI CM GA GN GW ML MR NE SN TD TG |
|
| 121 | Ep: the epo has been informed by wipo that ep was designated in this application | ||
| DFPE | Request for preliminary examination filed prior to expiration of 19th month from priority date (pct application filed before 20040101) | ||
| WWE | Wipo information: entry into national phase |
Ref document number: 65222/99 Country of ref document: AU |
|
| ENP | Entry into the national phase |
Ref document number: 2347736 Country of ref document: CA Ref country code: CA Ref document number: 2347736 Kind code of ref document: A Format of ref document f/p: F |
|
| ENP | Entry into the national phase |
Ref country code: JP Ref document number: 2000 577142 Kind code of ref document: A Format of ref document f/p: F |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 1999953250 Country of ref document: EP |
|
| REG | Reference to national code |
Ref country code: DE Ref legal event code: 8642 |
|
| AK | Designated states |
Kind code of ref document: A3 Designated state(s): AE AL AM AT AU AZ BA BB BG BR BY CA CH CN CR CU CZ DE DK DM EE ES FI GB GD GE GH GM HR HU ID IL IN IS JP KE KG KP KR KZ LC LK LR LS LT LU LV MD MG MK MN MW MX NO NZ PL PT RO RU SD SE SG SI SK SL TJ TM TR TT TZ UA UG UZ VN YU ZA ZW |
|
| AL | Designated countries for regional patents |
Kind code of ref document: A3 Designated state(s): GH GM KE LS MW SD SL SZ TZ UG ZW AM AZ BY KG KZ MD RU TJ TM AT BE CH CY DE DK ES FI FR GB GR IE IT LU MC NL PT SE BF BJ CF CG CI CM GA GN GW ML MR NE SN TD TG |
|
| WWP | Wipo information: published in national office |
Ref document number: 1999953250 Country of ref document: EP |
|
| WWG | Wipo information: grant in national office |
Ref document number: 65222/99 Country of ref document: AU |
|
| WWG | Wipo information: grant in national office |
Ref document number: 1999953250 Country of ref document: EP |