WO2000069835A2 - Process for manufacturing diazepine derivatives - Google Patents

Process for manufacturing diazepine derivatives Download PDF

Info

Publication number
WO2000069835A2
WO2000069835A2 PCT/EP2000/004033 EP0004033W WO0069835A2 WO 2000069835 A2 WO2000069835 A2 WO 2000069835A2 EP 0004033 W EP0004033 W EP 0004033W WO 0069835 A2 WO0069835 A2 WO 0069835A2
Authority
WO
WIPO (PCT)
Prior art keywords
chloro
methyl
benzodiazepine
dihydro
general formula
Prior art date
Legal status (The legal status is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the status listed.)
Ceased
Application number
PCT/EP2000/004033
Other languages
French (fr)
Other versions
WO2000069835A3 (en
Inventor
Fabienne Hoffmann-Emery
Current Assignee (The listed assignees may be inaccurate. Google has not performed a legal analysis and makes no representation or warranty as to the accuracy of the list.)
F Hoffmann La Roche AG
Original Assignee
F Hoffmann La Roche AG
Priority date (The priority date is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the date listed.)
Filing date
Publication date
Priority to MXPA01011445A priority Critical patent/MXPA01011445A/en
Application filed by F Hoffmann La Roche AG filed Critical F Hoffmann La Roche AG
Priority to CA002373506A priority patent/CA2373506C/en
Priority to DK00929501T priority patent/DK1202977T3/en
Priority to CN008074224A priority patent/CN101426771B/en
Priority to AT00929501T priority patent/ATE256669T1/en
Priority to JP2000618252A priority patent/JP3590590B2/en
Priority to EP00929501A priority patent/EP1202977B1/en
Priority to DE60007324T priority patent/DE60007324T2/en
Priority to AU47555/00A priority patent/AU4755500A/en
Publication of WO2000069835A2 publication Critical patent/WO2000069835A2/en
Anticipated expiration legal-status Critical
Publication of WO2000069835A3 publication Critical patent/WO2000069835A3/en
Ceased legal-status Critical Current

Links

Classifications

    • CCHEMISTRY; METALLURGY
    • C07ORGANIC CHEMISTRY
    • C07DHETEROCYCLIC COMPOUNDS
    • C07D243/00Heterocyclic compounds containing seven-membered rings having two nitrogen atoms as the only ring hetero atoms
    • C07D243/06Heterocyclic compounds containing seven-membered rings having two nitrogen atoms as the only ring hetero atoms having the nitrogen atoms in positions 1 and 4
    • C07D243/10Heterocyclic compounds containing seven-membered rings having two nitrogen atoms as the only ring hetero atoms having the nitrogen atoms in positions 1 and 4 condensed with carbocyclic rings or ring systems
    • C07D243/121,5-Benzodiazepines; Hydrogenated 1,5-benzodiazepines
    • CCHEMISTRY; METALLURGY
    • C07ORGANIC CHEMISTRY
    • C07DHETEROCYCLIC COMPOUNDS
    • C07D487/00Heterocyclic compounds containing nitrogen atoms as the only ring hetero atoms in the condensed system, not provided for by groups C07D451/00 - C07D477/00
    • C07D487/02Heterocyclic compounds containing nitrogen atoms as the only ring hetero atoms in the condensed system, not provided for by groups C07D451/00 - C07D477/00 in which the condensed system contains two hetero rings
    • C07D487/04Ortho-condensed systems
    • CCHEMISTRY; METALLURGY
    • C07ORGANIC CHEMISTRY
    • C07DHETEROCYCLIC COMPOUNDS
    • C07D243/00Heterocyclic compounds containing seven-membered rings having two nitrogen atoms as the only ring hetero atoms
    • C07D243/06Heterocyclic compounds containing seven-membered rings having two nitrogen atoms as the only ring hetero atoms having the nitrogen atoms in positions 1 and 4
    • C07D243/10Heterocyclic compounds containing seven-membered rings having two nitrogen atoms as the only ring hetero atoms having the nitrogen atoms in positions 1 and 4 condensed with carbocyclic rings or ring systems
    • C07D243/141,4-Benzodiazepines; Hydrogenated 1,4-benzodiazepines

Definitions

  • the compounds of general formula I are valuable intermediate products for the manufacture of imidazo[ l,5-a] [l,4]diazepine derivatives, like for instance 7-chloro-3-(5- dimethylaminomethyl-[ 1,2,4] oxadiazol-3-yl)-5-methyl-4,5-dihydro- imidazo[l,5-a] [ l,4]benzodiazepin-6-one, which diazepine derivatives show excellent psychopharmacological properties as agonists of the central benzodiazepine receptors.
  • the compounds of formula II can be obtained, on their turn, by reacting a compound of formula
  • the last step of the mentioned production pathway is characterised by low yields. This is mainly due to a low conversion of the reactants and, in certain cases, also to a low selectivity towards the desired product because of the formation of a side product of general formula
  • the brown powder obtained was digested in 2.0 ml water at 0°C for 1 hour, filtered off and washed with 2.0 ml cold water to yield, after drying under vacuum, 0.57 g (91% mol) (S)-6-chloro-l,2,3,l la- tetrahydro-5H-pyrrolo[2,l-c] [ l,4]-benzodiazepine-5,l l(10H)-dione as a beige powder of m.p. > 250°C.

Landscapes

  • Chemical & Material Sciences (AREA)
  • Organic Chemistry (AREA)
  • Pharmaceuticals Containing Other Organic And Inorganic Compounds (AREA)
  • Nitrogen Condensed Heterocyclic Rings (AREA)
  • Nitrogen And Oxygen Or Sulfur-Condensed Heterocyclic Ring Systems (AREA)

Abstract

The present invention relates to a process for manufacturing diazepine derivatives of general formula (I) wherein R1 is lower alkyl and R2 is hydrogen, or R?1 and R2¿ are together -(CH¿2?)n- and n is 2 or 3; R?3¿ is halogen, lower alkyl, lower alkoxy and m is 0, 1 or 2; R4 is hydrogen or lower alkyl. The compounds of general formula (I) are valuable intermediate products for the manufacture of imidazo[1,5-a][1,4]diazepine derivatives, like for instance 7-chloro-3-(5-dimethylaminomethyl-[1,2,4]oxadiazol-3-yl)-5-methyl-4,5-dihydro-imidazo[1,5-a][1,4]benzodiazepin-6-one, which diazepine derivatives show excellent psychopharmacological properties as agonists of the central benzodiazepine receptors.

Description

Process for manufacturing diazepine derivatives
The present invention relates to a process for manufacturing diazepine derivatives of the general formula
Figure imgf000003_0001
wherein R1 is lower alkyl and R2 is hydrogen, or R1 and R2 are together -(CH2)n- and n is 2 or 3; R3 is halogen, lower alkyl, lower alkoxy and m is 0, 1 or 2; R4 is hydrogen or lower alkyl.
The compounds of general formula I are valuable intermediate products for the manufacture of imidazo[ l,5-a] [l,4]diazepine derivatives, like for instance 7-chloro-3-(5- dimethylaminomethyl-[ 1,2,4] oxadiazol-3-yl)-5-methyl-4,5-dihydro- imidazo[l,5-a] [ l,4]benzodiazepin-6-one, which diazepine derivatives show excellent psychopharmacological properties as agonists of the central benzodiazepine receptors.
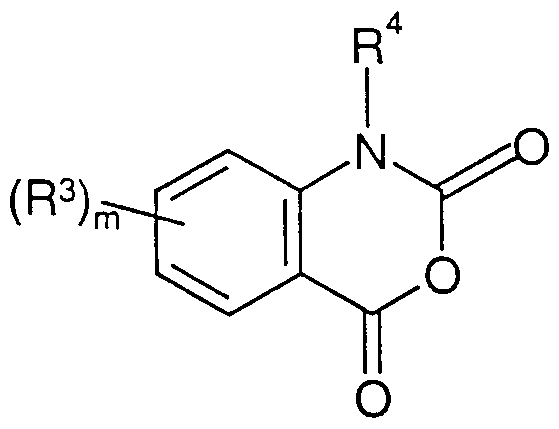
The compounds of general formula I are obtained by the known process consisting of reacting a compound of general formula
Figure imgf000003_0002
wherein R > 3' , r R>4 and m have the meaning mentioned above, with a compound of general formula
H— N-CH— COOH R FT wherein R and R have the meaning mentioned above. This reaction step takes place in a polar solvent such as for instance DMF, under atmospheric pressure and at a temperature between 110°C and the boiling point of the reaction mixture.
The compounds of formula II can be obtained, on their turn, by reacting a compound of formula
Figure imgf000004_0001
wherein R and m have the meaning mentioned above,
with: a) phosgene and hydrochloric acid in THF; or b) ethyl haloformiate, e.g. ethyl chloroformiate, in dioxane and subsequent treatment with acetylchloride.
Both steps take place in a batch system, under atmospheric pressure and at the boiling temperature of the reaction mixture (see e.g. G. M. Coppola, "The Chemistry of Isatoic Anhydride", Synthesis, Georg Thieme Verlag, (1980), pp 505-535).
The last step of the mentioned production pathway is characterised by low yields. This is mainly due to a low conversion of the reactants and, in certain cases, also to a low selectivity towards the desired product because of the formation of a side product of general formula
Figure imgf000004_0002
These low yield and selectivity imply higher costs for the production of the compounds of formula I and lead to important disposal problems since the compounds of formula V cannot be used for other purposes and must be therefore destroyed or recycled.
The problem at the root of the present invention is therefore to provide a process for manufacturing the compounds of general formula I which can overcome the disadvantages mentioned above.
The problem is solved, according to the present invention, by a process for manufacturing diazepine derivatives of the general formula I, comprising the step of reacting a compound of general formula II with a compound of general formula III, characterised in that said compound of general formula II and said compound of general formula III undergo chemical reaction in the absence of a solvent or in the presence of an apolar solvent.
It has been surprisingly found that the conversion, and in certain cases also the selectivity, towards the compound of formula I strongly increases if the reaction components (i.e. compounds of formula II and III) are not solvated in the reaction mixture. This situation can take place only if no solvent at all is added to the reaction mixture or if the reactants and/or products are not soluble in a given solvent. Being the present compounds of polar nature, apolar solvents can be used in the process of the invention for achieving the wished results.
Particularly preferred solvents are substituted benzene rings, such as xylenes, mesitylene, ethylbenzene, isopropylbenzene, etc. Most preferably, p-xylene or a mixture of xylenes are used as solvent for carrying out the process according to the present invention.
The reaction temperature is preferably set from 0 to 30°C under the boiling temperature of the reaction mixture.
The process of the present invention is particularly suitable for the manufacture of 6- Chloro-3,4-dihydro-4-methyl-2H-l,4-benzodiazepine-2,5(lH)-dione.
By way of examples, preferred embodiments of the present invention will now be described. Comparative tests were made in which the compounds of formulae II and III underwent reaction in the conventional manner, i.e. using DMF as (polar) solvent. The yields of compound I (Y(CI)) depicted in Table 1 were measured on the purified product. The ratios of compounds I and V depicted in Table 2 (R(CI) and R(CV)) were directly obtained from the HPLC measurements (HP1050, column CC70/4 nucleosil 100- 5C18HD), and refer to the molar percentage of Cl and CV in the crude product of the reaction.
Y(CI) = 100* [mol Cl/mol CII] Cl, CII, CV = compound of general formula I, II, V
Example 1
6-Chloro-3,4-dihydro-4-methyl-2H-l,4-benzodiazepine-2,5(lH)-dione.
25.0 g (126 mMol) 5-chloro-lH-benzo[d] [l,3]xazin-2,4-dione and 12.4 g (139 mMol) sarcosine were suspended in 100 ml p-xylene and heated at reflux (oil bath temperature (Text) <150°C) for 2 hours. After cooling to room temperature (r.t.), the suspension was stirred one more hour. The precipitate was filtered off, washed with 25 ml p-xylene twice and dried at 50°C under vacuum. The solid obtained was digested in 75 ml water one hour at 0°C, filtered off, washed with 25 ml water and dried under vacuum for 18 hours at 80°C to yield 25.2 g (88% mol) of 6-chloro-3,4-dihydro-4-methyl-2H-l,4-benzodiazepine- 2,5(lH)-dione of m.p. = 230-232°C.
MS (El): 224 (M +, 52); 153 (68); 44(100).
Example 2
6-Methyl-3,4-dihydro-4-methyl-2H-l,4-benzodiazepine-2,5(lH)-dione.
1.0 g (5.6 mMol) 5-methyl-lH-benzo[d] [l,3]xazin-2,4-dione and 0.57 g (6.4 mMol) sarcosine were suspended in 4 ml p-xylene and heated at reflux (Text<150oC) for 5.5 hours. p-Xylene was removed under reduced pressure and the solid residue was digested in 5.0 ml water one hour at 0°C, filtered off, and dried under vacuum for 18 hours at 80°C to yield 0.93 g (81% mol) of 6-methyl-3,4-dihydro-4-methyl-2H-l,4-benzodiazepine-2,5(lH)- dione of m.p. = 203.5-205°C. MS (El): 204 (M\ 94); 175 (38); 133 (100); 44 (100).
Example 3
7-Fluoro-3,4-dihydro-4-methyl-2H-l,4-benzodiazepine-2,5(lH)-dione.
1.0 g (5.5 mMol) 6-fluoro-lH-benzo[d] [l,3]xazin-2,4-dione and 0.54 g (6.1 mMol) sarcosine were suspended in 4.0 ml p-xylene and heated to reflux for 4 hours. The suspension was cooled to r.t. and the precipitate filtered off. The solid obtained was digested 30 minutes at 0°C in 5 ml deionised water, filtered off and dried for 16 hours at 60°C under vacuum to yield 0.92 g (80% mol) of 7-fluoro-3,4-dihydro-4-methyl-2H-l,4- benzodiazepine-2,5(lH)-dione of m.p. > 250°C.
MS (El): 208 (M \ 94); 179 (100); 137(92).
Example 4
7-Chloro-6-fluoro-3,4-dihydro-4-methyl-2H-l,4-benzodiazepine-2,5(lH)-dione.
1.0 g (4.6 mMol) 5-chloro-6-fluoro-lH-benzo[d] [l,3]xazin-2,4-dione and 0.45 g (5.0 mMol) sarcosine were suspended in 4.0 ml p-xylene and heated to reflux (Text = 145°C) for 7 hours. Solvent was removed under reduced pressure and the residue was digested in 2.0 ml deionised water 1 hour at r.t.. The precipitate was filtered off and crystallized from 10 ml methanol and 10 ml diethylether to give 0.63 g (56% mol) of 7-chloro-6-fluoro-3,4- dihydro-4-methyl-2H-l,4-benzodiazepine-2,5(lH)-dione of m.p. > 250°C.
Concentration of the mother liquors and crystallization from 3 ml methanol and 9 ml diethylether gave an additional 0.13 g (11%) of 7-chloro-6-fluoro-3,4-dihydro-4-methyl- 2H-l,4-benzodiazepine-2,5(lH)-dione of m.p. > 250°C.
MS (El): 242 (M'+, 56); 213 (58); 171(76); 44 (100). Example 5
(S)-6-Chloro-l,2,3,lla-tetrahydro-5H-pyτrolo[2,l-c] [l,4]-benzodiazepine-5,ll(10H)- dione.
0.50 g (2.5 mMol) 5-chloro-lH-benzo[d] [ l,3]xazin-2,4-dione and 0.32 g (2.8 mMol) L- proline were suspended in 4.0 ml p-xylene and heated at reflux (Text<150°C) for 2.5 hours (gives a yellow solution). Upon cooling to r.t., a precipitate formed which was filtered off and dried at 60°C under vacuum. The solid obtained was digested in 1.5 ml water one hour at 0°C, filtered off, washed with 1.0 ml water and dried under vacuum for 16 hours at 60°C to give 0.49 g (78% mol) of (S)-6-chloro- l)2,3,lla-tetrahydro-5H-pyrrolo[2,l-c] [ l)4]- benzodiazepine-5,l l(10H)-dione of m.p. > 250°C.
MS (El): 250 (M +, 40); 221 (30); 70( 100).
Example 6
(S)-l,10a-5-Chloro-2H-azeto[2,l-c] [ l,4]benzodiazepine-4,10(9H)-dione.
1.0 g (5.1 mMol) 5-chloro-lH-benzo[d] [ l,3]xazin-2,4-dione and 0.56 g (5.6 mMol) (S)- azetidine-2-carboxylic acid were suspended in 6.0 ml p-xylene and heated to reflux for 24 hours. p-Xylene was removed under reduced pressure and the residue was partitioned between dichloromethane and water and the aqueous phase extracted with dichloromethane. The combined organic extracts were dried (Na2SO4) and the solvent removed under reduced pressure. The brown solid obtained was digested in 5 ml tert- butyl-methylether for 16 hours at r.t, filtered and dried under reduced pressure to give 0.99 g (82% mol) of (S)-l,10a-5-chloro-2H-azeto[2,l-c] [ l,4]benzodiazepine-4,10(9H)- dione as a beige powder of m. p. = 180-198°C
MS (El): 236 (M \ 44); 180 (24); 153(62), 56 ( 100).
Example 7
(S)- 1-Methyl- 1 ,2,3, 11 a-tetrahydro-5H-pyrrolo [2,1-c] [ 1 ,4] -benzodiazepine-5, 11 ( 10H)- dione.
0.5 g (2.82 mMol) N-metyl-lH-benzo[d] [l,3]xazin-2,4-dione and 0.36 g (3.1 mMol) L- proline were suspended in 1.0 ml p-xylene and heated to reflux for 1 hour (goes into solution upon heating). After cooling to r.t., the reaction mixture was diluted with 10 ml dichloromethane and 5 ml deionised water and the phases separated. The aqueous phase was extracted with 8 ml dichloromethane twice. The combined organic extracts were dried (Na2SO ) and evaporated. The residue was digested in 2 ml fert-butyl-methylether for 2 hours at r.t. to give 0.53 g (81.5% mol) of (S)-l-methyl- 1,2,3,1 la-tetrahydro-5H- pyrrolo[2,l-c] [l,4]-benzodiazepine-5,l l(10H)-dione as beige crystals of m. p. = 117- 118.5°C.
MS (El): 230 (M +, 56); 161 (99); 133 (90); 105 (88); 70(100).
Example 8 6-Chloro-3,4-dihydro-4-methyl-2H-l,4-benzodiazepine-2,5(lH)-dione.
1.0 g (5.0 mMol) 5-chloro-lH-benzo[d] [l,3]xazin-2,4-dione and 0.50 g (5.56 mMol) sarcosine were suspended in 4.0 ml of a mixture of xylenes and heated at reflux (Teχt<150°C) for 4 hours. After cooling to room temperature, the suspension was stirred one more hour. The precipitate was filtered off, washed with 1.5 ml hexane twice and dried at 60°C under vacuum. The solid obtained was digested in 3.0 ml water one hour at 0°C, filtered off, washed with 2.0 ml water and dried under vacuum for 4 hours at 60°C to yield 0.86 g (74% mol) of 6-chloro-3,4-dihydro-4-methyl-2H-l,4-benzodiazepine-2,5(lH)- dione of m.p. = 235-237°C.
MS (El): 224 (M \48); 195(34); 153(60) 126(36), 44(100).
Example 9 7-Chloro-3,4-dihydro-4-methyl-2H-l,4-benzodiazepine-2,5(lH)-dione.
1.0 g (5.06 mMol) 6-chloro-lH-benzo[d] [l,3]xazin-2,4-dione and 0.67 g (7.59 mMol) sarcosine were thoroughly mixed and heated to 140°C for 2 hours then 150°C for 20 hours. The brown powder obtained was cooled to r.t. and digested in 4.0 ml water at 0°C for 1 hour, filtered and washed with 1.0 ml water. After drying under vacuum, 1.0 g (88% mol) 7-chloro-3,4-dihydro-4-methyl-2H-l,4-benzodiazepine-2,5(lH)-dione was obtained as a beige powder of m. p. > 250°C.
MS (El): 224 (M \78); 195 (86); 153(80), 44(100). Example 10
(S)-6-Chloro-l,2,3,lla-tetrahydro-5H-pyrrolo[2)l-c] [l,4]-benzodiazepine-5,ll(10H)- dione.
0.50 g (2.5 mMol) 5-chloro-lH-benzo[d] [l,3]xazin-2,4-dione and 0.43 g (3.75 mMol) L- proline were finely ground together and heated to 150°C for 18 hours. The brown powder obtained was digested in 2.0 ml water at 0°C for 1 hour, filtered off and washed with 2.0 ml cold water to yield, after drying under vacuum, 0.57 g (91% mol) (S)-6-chloro-l,2,3,l la- tetrahydro-5H-pyrrolo[2,l-c] [ l,4]-benzodiazepine-5,l l(10H)-dione as a beige powder of m.p. > 250°C.
MS (El): 250 (M +,36); 221 (28); 194(28); 153(32); 126(30); 70(100).
Table 1: Reaction yields after purification.
Figure imgf000010_0001
Table 2: Ratios of Cl and CV in the crude product
Figure imgf000010_0002
As showed in the above tables, the process according to the present invention leads to yields in the desired product which are much higher than those obtainable with conventional processes. Therefore, the process according to the present invention enables an increase of the productivity, thereby decreasing costs and disposal problems.
As stated above, the products obtained with the process according to the invention can be used for manufacturing imidazo [ 1 ,5-a] [ 1 ,4] diazepine derivatives with excellent psychopharmacological properties. Example 11 illustrates a possible method for producing one of such diazepine derivatives.
Example 11 7-chloro-3-(5-dimethylaminomethyl- [1,2,4] oxadiazol-3-yl)-5-methyl-4,5-dihydro- imidazo[l,5-a] [l,4]benzodiazepin-6-one.
Ethyl 7-chloro-5,6-dihydro-5-methyl-6-oxo-4H-imidazo[l,5-a][l,4]benzodiazepine- 3- carboxylate.
25.0 g 6-Chloro-3,4-dihydro-4-methyl-2H-l,4-benzodiazepine-2,5( lH)-dione were suspended under stirring and argon atmosphere in 200 ml toluene and 32.1 ml N,N- dimethyl-p-toluidine. The suspension was heated to 100°C and 11.2 ml phosphorus oxychloride were added over 30 minutes and stirring was pursued two and an half hours at 100°C. The dark-orange solution was cooled to 40°C and toluene was removed under reduced pressure to give 82 g of a dark-orange oil.
Meanwhile, 81.2 ml hexamethyldisilazane and 265 ml tetrahydrofuran were mixed and cooled to -35°C. 229.5 ml Butyllithium were added over 45 minutes and, after stirring 30 minutes at -35°C, a solution of 35.2 g ethyl(dimethylamino-methylenamino)acetate in 70.4 ml tetrahydrofuran was added over 30 minutes. The orange solution obtained was stirred one more hour at -35°C and a solution of the crude iminochloride in 100 ml tetrahydrofuran was added over 1 hour at -15°C. The dark red solution was stirred one hour at -15°C, then 18 hours at room temperature (r.t.). 75 ml Acetic acid were added in 10 minutes, then 75 ml deionized water were added in one portion and the orange suspension was heated at reflux for two hours. Tetrahydrofuran was removed under reduced pressure and the residue was partitioned between 200 ml dichloromethane and 100 ml deionized water. The phases were separated and the organic phase was washed with 100 ml aqueous HC1 IN twice and with 100 ml deionized water. The aqueous phases were extracted twice with 100 ml dichloromethane. The combined organic extracts were dried (Na2SO ) and evaporated. The residue was digested in 200 ml n-heptane 30 minutes at r.t. and filtered off. The sticky crystals obtained were digested at reflux for 30 minutes in 213.5 ml efhanol, then stirred 3 hours to r.t. and 2 hours at -20°C. The precipitate (ethyl 7- chloro-5,6-dihydro-5-methyl-6-oxo-4H-imidazo[l,5-a] [l,4]benzodiazepine-3- carboxylate) was filtered off, washed three times with 20 ml ethanol and dried under reduced pressure 16 hours at 60°C. Crude product: 23.4 g as a beige powder, m.p. 225.5- 226.5 °C
7-Chloro-5,6-dihydro-5-methyl-6-oxo-4H-imidazo[l,5-a][l,4]benzodiazepine-3- carboxamide.
22.8 g Ethyl 7-chloro-5,6-dihydro-5-methyl-6-oxo-4H-imidazo[l,5-a] [l,4]- benzodiazepine-3-carboxylate were suspended under stirring and argon atmosphere in 91.2 ml 1,4-dioxane. 14.1 ml Formamide and 13.9 ml sodium methanolate were successively added to yield a clear light-orange solution, which turned to a white suspension after 10 minutes. This suspension was stirred two hours at 30°C. 200 ml Deionized water were added in one portion and 1,4-dioxane was distilled off at 40°C under reduced pressure. The remaining white suspension was stirred two hours at 0°C and filtered. The precipitate (7-chloro-5,6-dihydro-5-methyl-6-oxo-4H- imidazo[ l,5-a] [ l,4]benzodiazepine-3-carboxamide) was washed with 50 ml deionized water three times and dried under reduced pressure for 18 hours at 80°C. Crude product:19.43 g as a white powder. m.p.>250°C
7-Chloro-5,6-dihydro-5-methyl-6-oxo-4H-imidazo[l,5-a][l,4]benzodiazepine-3-carbonitrile.
19.0 g 7-Chloro-5,6-dihydro-5-methyl-6-oxo-4H-imidazo[ l,5-a] [l,4]benzodiazepine-3- carboxamide were suspended under stirring and argon atmosphere in 95 ml 1,4-dioxane and 6.58 ml phosphorous oxychloride were added in one portion. The reaction mixture was heated to reflux for one hour giving a yellow solution, which was concentrated at 50°C under reduced pressure. The residue was digested in 100 ml deionized water for two hours at r.t.. The precipitate (7-chloro-5,6-dihydro-5-methyl-6-oxo-4H- imidazo[ l,5-a] [ l,4]benzodiazepine-3-carbonitrile) was filtered off, washed three times with 30 ml deionized water and dried under vacuum at 80°C for 18 hours. Crude product:
17.3 g as a light yellow powder, m.p. 238.5-239.5°C
7-Chloro-5,6-dihydro-5-methyl-6-oxo-4H-imidazo[l,5-a][l,4]benzodiazepine-3- carboxamidoxime.
16.8 g 7-Chloro-5,6-dihydro-5-methyl-6-oxo-4H-imidazo[l,5-a] [ l,4]benzodiazepine-3- carbonitrile were suspended under stirring and argon atmosphere in 101 ml N,N- dimethylformamide and 13.48 g hydroxylamine hydrochloride were added in one portion. 34.2 ml Sodium methanolate were then added over 60 minutes to the yellow suspension, which turned to a colorless suspension. It was stirred one more hour at r.t., then cooled to 0-2°C and 202 ml deionized water were added over 30 minutes. After stirring one more hour at 0°C, the precipitate (7-chloro-5,6-dihydro-5-methyl-6-oxo-4H- imidazo[l,5-a] [l,4]benzodiazepine-3-carboxamidoxime (VIII)) was filtered off, washed twice with 40 ml deionized water and dried under vacuum at 70°C for 18 hours. Crude product: 17.84 g as a white powder. m.p.>250°C
7-Chloro-3-(5-chloromethyl- [ 1,2,4] oxadi zol-3-yl)-5-methyl-4, 5-dϊhydro- imidazo[l,5-a][l,4]benzodiazepin-6-one.
8.0 g 7-Chloro-5,6-dihydro-5-methyl-6-oxo-4H-imidazo[ l,5-a] [l,4]benzodiazepine-3- carboxamidoxime and 1.0 g magnesium oxide were suspended under stirring and argon atmosphere in 160 ml 1,4-dioxane. 2.7 ml Chloracetyl chloride were added in one portion and the white thick gel obtained was stirred 4 hours at r.t. and then 17 hours at reflux to give a lightly orange fluid suspension. 100 ml Dioxane were distilled off and the reaction mixture was cooled to room temperature. 180 ml Deionized water were added within 15 minutes and the suspension was stirred 1 hour at r.t.. The precipitate was filtered off, washed with 50 ml deionized water twice and dried under vacuum at 80°C for 18 hours. Crude product: 8.3 g as a light pink powder. This crude product was dissolved in 120 ml tetrahydrofuran at reflux and 0.83 g active charcoal Darco G 60 were added. The system was refluxed 1 hour, then filtered on 25 g Dicalit-Speedex and the filter cake was washed with three portions of 50 ml warm tetrahydrofuran. The filtrate was concentrated at 40°C under reduced pressure. The residue was digested in 80 ml ethanol 1 hour at reflux, then stirred 16 hours at r.t. and finally 2 hours at 2°C. The precipitate (7-chloro-3-(5- chloromethyl- [ 1,2,4] oxadiazol-3-yl)-5-methyl-4,5-dihydro-imidazo [l,5-a] [l,4]benzo- diazepin-6-one (IX)) was filtered off, washed with 2 portions of 25 ml cold tert-butyl methyl- ether and dried under vacuum 5 hours at 80°C. Crude product: 7.6 g as a light beige powder, m.p. 234-238°C
7-Chloro-3-(5-dimethylaminomethyl-[ 1,2,4] oxadiazol-3-yl)-5-methyl-4,5-dihydro- imidazo[l,5-a][l,4]benzodiazepin-6-one.
7.0 g 7-Chloro-3-(5-chloromethyl- [1,2,4] oxadiazol-3-yl)-5-methyl-4,5-dihydro-imidazo- [1,5-a] [l,4]benzodiazepin-6-one were suspended under stirring and argon atmosphere in 70 ml 1,4-dioxane and 25.7 ml dimethylamine (33% in ethanol) were added over 60 minutes. The reaction mixture was stirred one more hour at r.t. and then the solvents were removed under reduced pressure at 35°C. The residue was partitioned between
50 ml dichloromethane and 20 ml deionized water. The phases were separated and the organic phase was washed twice with 20 ml deionized water. The aqueous phases were extracted separately with the same portion of 25 ml dichloromethane, twice. The combined organic extracts were dried (Na2SO ) and the solvent was removed under reduced pressure. Crude product: 8.0 g as a light yellow foam.
Purification The crude product was dissolved in 40 ml ethanol at reflux and 400 mg active charcoal Darco G 60 were added. The system was stirred 1 hour at reflux, then filtered on a hot pad of Dicalit Speedex, which was washed with two portions of 40 ml hot ethanol. The filtrate was concentrated to 14 g under reduced pressure, heated to reflux and at this temperature and 40 ml terr-butyl-methylether were added over 5 minutes. The suspension was cooled slowly to r.t., stirred 16 hours, further cooled to 2°C. After stirring 1 hour at 2°C, the precipitate was filtered off, washed with 20 ml ter -butyl-methylether and dried 1 hour at 60°C under vacuum. The so obtained powder was dissolved at reflux in 26 ml ethyl acetate. 6.5 ml Ethyl acetate were then distilled off and the turbid solution obtained was slowly cooled to r.t., then to 0°C. After 1 hour stirring at 0°C, the precipitate was filtered off, washed with 10 ml cold tert-butyl-methylether and dried under vacuum at 60°C for 16 hours. The so obtained powder (7-chloro-3-(5-dimethylaminomethyl-[l,2,4]oxadiazol-3- yl)-5-methyl-4,5-dihydro-imidazo[l,5-a] [l,4]benzodiazepin-6-one (I)) was crystallized a second time in 24.3 ml ethyl acetate according to the procedure described above. Product: 5.5 g as a white powder, m.p. 151.5-153°C.

Claims

Claims
1. Process for manufacturing diazepine derivatives of the general formula I
Ff
I
O R
wherein R1 is lower alkyl and R2 is hydrogen, or R1 and R2 are together -(CH )n- and n is 2 or 3; R" is halogen, lower alkyl, lower alkoxy and m is 0, 1 or 2; R4 is hydrogen or lower alkyl, comprising the step of reacting a compound of general formula
Figure imgf000015_0001
with a compound of general formula
H — N-CH — COOH
' 1 ' 2 R R
wherein Rl, R2, R3, R4, m and n have the meanings mentioned above, characterised in that said compound of general formula II and said compound of general formula III undergo chemical reaction in the absence of a solvent or in the presence of an apolar solvent.
2. The process according to claim 1, wherein the solvent is a substituted benzene ring.
3. The process according to claim 2, wherein the solvent is p-xylene.
4. The process according to claim 2, wherein the solvent is a mixture of xylenes.
5. The process according to any one of claims 1 to 4, wherein the compound of formula I is 6-Chloro-3,4-dihydro-4-methyl-2H-l,4-benzodiazepine-2,5(lH)-dione.
6. The process according to any one of claims 1 to 5, wherein the reaction temperature is from 0 to 30°C under the boiling temperature of the reaction mixture.
7. Product obtained by the process according to any one of claims 1 to 6.
8. The invention as hereinbefore described.
PCT/EP2000/004033 1999-05-12 2000-05-05 Process for manufacturing diazepine derivatives Ceased WO2000069835A2 (en)

Priority Applications (9)

Application Number Priority Date Filing Date Title
DE60007324T DE60007324T2 (en) 1999-05-12 2000-05-05 METHOD FOR PRODUCING DIAZEPINE DERIVATIVES
CA002373506A CA2373506C (en) 1999-05-12 2000-05-05 Process for manufacturing diazepine derivatives
DK00929501T DK1202977T3 (en) 1999-05-12 2000-05-05 Process for the preparation of diazepine derivatives
CN008074224A CN101426771B (en) 1999-05-12 2000-05-05 Process for manufacturing diazepine derivatives
AT00929501T ATE256669T1 (en) 1999-05-12 2000-05-05 METHOD FOR PRODUCING DIAZEPINE DERIVATIVES
MXPA01011445A MXPA01011445A (en) 1999-05-12 2000-05-05 Process for manufacturing diazepine derivatives.
EP00929501A EP1202977B1 (en) 1999-05-12 2000-05-05 Process for manufacturing diazepine derivatives
JP2000618252A JP3590590B2 (en) 1999-05-12 2000-05-05 Diazepine manufacturing method
AU47555/00A AU4755500A (en) 1999-05-12 2000-05-05 Process for manufacturing diazepine derivatives

Applications Claiming Priority (2)

Application Number Priority Date Filing Date Title
EP99109514 1999-05-12
EP99109514.2 1999-05-12

Publications (2)

Publication Number Publication Date
WO2000069835A2 true WO2000069835A2 (en) 2000-11-23
WO2000069835A3 WO2000069835A3 (en) 2002-03-07

Family

ID=8238166

Family Applications (1)

Application Number Title Priority Date Filing Date
PCT/EP2000/004033 Ceased WO2000069835A2 (en) 1999-05-12 2000-05-05 Process for manufacturing diazepine derivatives

Country Status (14)

Country Link
US (1) US6281353B1 (en)
EP (1) EP1202977B1 (en)
JP (1) JP3590590B2 (en)
KR (1) KR100469813B1 (en)
CN (1) CN101426771B (en)
AT (1) ATE256669T1 (en)
AU (1) AU4755500A (en)
CA (1) CA2373506C (en)
DE (1) DE60007324T2 (en)
DK (1) DK1202977T3 (en)
ES (1) ES2209880T3 (en)
JO (1) JO2259B1 (en)
MX (1) MXPA01011445A (en)
WO (1) WO2000069835A2 (en)

Families Citing this family (5)

* Cited by examiner, † Cited by third party
Publication number Priority date Publication date Assignee Title
US20100234359A1 (en) * 2007-08-20 2010-09-16 Evotec Neurosciences Gmbh Treatment of sleep disorders
US20090054412A1 (en) * 2007-08-20 2009-02-26 John Alan Kemp Treatment of Sleep Disorders
CN109134471B (en) * 2017-06-27 2020-05-15 浙江京新药业股份有限公司 Solid state form of benzodiazepine compound and preparation method and application thereof
CN111620834B (en) * 2019-10-15 2022-05-24 浙江京新药业股份有限公司 A method for preparing benzodiazepine compound and its intermediate
CN113045574B (en) * 2019-12-26 2024-06-07 浙江京新药业股份有限公司 Process for preparing diazapine derivatives and intermediates therefor

Family Cites Families (4)

* Cited by examiner, † Cited by third party
Publication number Priority date Publication date Assignee Title
CA1185602A (en) * 1981-02-27 1985-04-16 Emilio Kyburz Imidazodiazepines
CA1184175A (en) 1981-02-27 1985-03-19 Walter Hunkeler Imidazodiazepines
CA1174673A (en) * 1981-02-27 1984-09-18 Walter Hunkeler Imidazodiazepines
US4352817A (en) * 1981-02-27 1982-10-05 Hoffmann-La Roche Inc. Imidazo-diazepines and their use

Also Published As

Publication number Publication date
CN101426771A (en) 2009-05-06
CN101426771B (en) 2011-11-16
CA2373506A1 (en) 2000-11-23
AU4755500A (en) 2000-12-05
JP3590590B2 (en) 2004-11-17
JO2259B1 (en) 2004-10-07
EP1202977B1 (en) 2003-12-17
MXPA01011445A (en) 2002-06-04
DE60007324D1 (en) 2004-01-29
WO2000069835A3 (en) 2002-03-07
KR100469813B1 (en) 2005-02-02
CA2373506C (en) 2007-09-11
DK1202977T3 (en) 2004-04-05
KR20010112485A (en) 2001-12-20
ATE256669T1 (en) 2004-01-15
EP1202977A2 (en) 2002-05-08
ES2209880T3 (en) 2004-07-01
DE60007324T2 (en) 2004-10-21
JP2002544265A (en) 2002-12-24
US6281353B1 (en) 2001-08-28

Similar Documents

Publication Publication Date Title
RU2145606C1 (en) Tricyclic dicarbonyl derivatives and drug based on said
CA2143246C (en) Imidazodiazepines
EP1149102B1 (en) Triazolo-pyridazine derivatives as ligands for gaba receptors
CA1067491A (en) Diazepine derivatives
JPH02258785A (en) New annulated indolo (3,2-c) lactam, preparation thereof, drug composition containing same as effective component, and preparation of said composition
NZ508290A (en) A substituted triazolo-pyridazine derivative, pharmaceutical compostions made therefrom
EP1202977B1 (en) Process for manufacturing diazepine derivatives
JP2001503759A (en) 3-carboxamide derivative of 5H-pyrrolo [2,1-c] [1,4] benzodiazepine
Jiang et al. α-Cyclization of tertiary amines. Part 1
WO2000047582A1 (en) Triazolo-pyridazine derivatives as ligands for gaba receptors
Podanyi et al. Nitrogen bridgehead compounds. Part 63. Ring-chain tautomerism of [(alpha-N-hetaryl) aminomethylene] malononitriles
SU725563A3 (en) Method of preparing imidazo-(1,5-a)(1,4) diazepine derivatives or their salts
US4075202A (en) S-triazolo-1,5-benzodiazpine-4-ones
MXPA97003288A (en) Oxazolil- and tiazolil imidazo-benzo- and tienodiazepinas and its use as a medicamen
US3575976A (en) 3-amino-1,2,8,9-tetraazaphenalenes
US3560500A (en) 3-halo-1,2,8,9-tetraazaphenalenes
FI67849C (en) DETERMINATION OF THERAPEUTIC THERAPEUTIC MEASURE I 3-STATIONARY SUBSTITUTE IMIDAZO (1,5-A) (1,4) DIAZEPINFOERENINGAR SAMT
NZ227983A (en) 5 cyclopropyl-3-isocyanomethyl-1,2,4-oxidiazole
IL49503A (en) Process for the preparation of 6-phenyl-4h-s-triazolo(4,3-a)(1,4)-benzodiazepines and intermediate 2-(3-(phthalimidomethyl)-4h-1,2,4-triazol-4-yl)benzophenones

Legal Events

Date Code Title Description
WWE Wipo information: entry into national phase

Ref document number: 00807422.4

Country of ref document: CN

AK Designated states

Kind code of ref document: A2

Designated state(s): AE AL AM AT AU AZ BA BB BG BR BY CA CH CN CU CZ DE DK EE ES FI GB GD GE GH GM HR HU ID IL IN IS JP KE KG KP KR KZ LC LK LR LS LT LU LV MA MD MG MK MN MW MX NO NZ PL PT RO RU SD SE SG SI SK SL TJ TM TR TT UA UG UZ VN YU ZA ZW

AL Designated countries for regional patents

Kind code of ref document: A2

Designated state(s): GH GM KE LS MW SD SL SZ TZ UG ZW AM AZ BY KG KZ MD RU TJ TM AT BE CH CY DE DK ES FI FR GB GR IE IT LU MC NL PT SE BF BJ CF CG CI CM GA GN GW ML MR NE SN TD TG

DFPE Request for preliminary examination filed prior to expiration of 19th month from priority date (pct application filed before 20040101)
121 Ep: the epo has been informed by wipo that ep was designated in this application
WWE Wipo information: entry into national phase

Ref document number: 2000929501

Country of ref document: EP

WWE Wipo information: entry into national phase

Ref document number: IN/PCT/2001/1536/CHE

Country of ref document: IN

ENP Entry into the national phase

Ref document number: 2373506

Country of ref document: CA

Ref document number: 2373506

Country of ref document: CA

Kind code of ref document: A

Ref document number: 2000 618252

Country of ref document: JP

Kind code of ref document: A

WWE Wipo information: entry into national phase

Ref document number: PA/a/2001/011445

Country of ref document: MX

WWE Wipo information: entry into national phase

Ref document number: 1020017014340

Country of ref document: KR

WWP Wipo information: published in national office

Ref document number: 1020017014340

Country of ref document: KR

AK Designated states

Kind code of ref document: A3

Designated state(s): AE AL AM AT AU AZ BA BB BG BR BY CA CH CN CU CZ DE DK EE ES FI GB GD GE GH GM HR HU ID IL IN IS JP KE KG KP KR KZ LC LK LR LS LT LU LV MA MD MG MK MN MW MX NO NZ PL PT RO RU SD SE SG SI SK SL TJ TM TR TT UA UG UZ VN YU ZA ZW

AL Designated countries for regional patents

Kind code of ref document: A3

Designated state(s): GH GM KE LS MW SD SL SZ TZ UG ZW AM AZ BY KG KZ MD RU TJ TM AT BE CH CY DE DK ES FI FR GB GR IE IT LU MC NL PT SE BF BJ CF CG CI CM GA GN GW ML MR NE SN TD TG

REG Reference to national code

Ref country code: DE

Ref legal event code: 8642

WWP Wipo information: published in national office

Ref document number: 2000929501

Country of ref document: EP

WWG Wipo information: grant in national office

Ref document number: 2000929501

Country of ref document: EP

WWG Wipo information: grant in national office

Ref document number: 1020017014340

Country of ref document: KR