WO2003097060A1 - Oral therapeutic or preventive drugs for pollakiuria and urinary incontinence or oral sleep inducers, containing tropolone derivatives - Google Patents
Oral therapeutic or preventive drugs for pollakiuria and urinary incontinence or oral sleep inducers, containing tropolone derivatives Download PDFInfo
- Publication number
- WO2003097060A1 WO2003097060A1 PCT/JP2003/006003 JP0306003W WO03097060A1 WO 2003097060 A1 WO2003097060 A1 WO 2003097060A1 JP 0306003 W JP0306003 W JP 0306003W WO 03097060 A1 WO03097060 A1 WO 03097060A1
- Authority
- WO
- WIPO (PCT)
- Prior art keywords
- isopropyl
- pyrimidyl
- piperazinomethyl
- urinary incontinence
- pollakiuria
- Prior art date
- Legal status (The legal status is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the status listed.)
- Ceased
Links
Classifications
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61K—PREPARATIONS FOR MEDICAL, DENTAL OR TOILETRY PURPOSES
- A61K31/00—Medicinal preparations containing organic active ingredients
- A61K31/33—Heterocyclic compounds
- A61K31/395—Heterocyclic compounds having nitrogen as a ring hetero atom, e.g. guanethidine or rifamycins
- A61K31/495—Heterocyclic compounds having nitrogen as a ring hetero atom, e.g. guanethidine or rifamycins having six-membered rings with two or more nitrogen atoms as the only ring heteroatoms, e.g. piperazine or tetrazines
- A61K31/496—Non-condensed piperazines containing further heterocyclic rings, e.g. rifampin, thiothixene or sparfloxacin
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61K—PREPARATIONS FOR MEDICAL, DENTAL OR TOILETRY PURPOSES
- A61K31/00—Medicinal preparations containing organic active ingredients
- A61K31/33—Heterocyclic compounds
- A61K31/395—Heterocyclic compounds having nitrogen as a ring hetero atom, e.g. guanethidine or rifamycins
- A61K31/495—Heterocyclic compounds having nitrogen as a ring hetero atom, e.g. guanethidine or rifamycins having six-membered rings with two or more nitrogen atoms as the only ring heteroatoms, e.g. piperazine or tetrazines
- A61K31/505—Pyrimidines; Hydrogenated pyrimidines, e.g. trimethoprim
- A61K31/506—Pyrimidines; Hydrogenated pyrimidines, e.g. trimethoprim not condensed and containing further heterocyclic rings
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P13/00—Drugs for disorders of the urinary system
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P13/00—Drugs for disorders of the urinary system
- A61P13/02—Drugs for disorders of the urinary system of urine or of the urinary tract, e.g. urine acidifiers
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P13/00—Drugs for disorders of the urinary system
- A61P13/10—Drugs for disorders of the urinary system of the bladder
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P25/00—Drugs for disorders of the nervous system
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P25/00—Drugs for disorders of the nervous system
- A61P25/20—Hypnotics; Sedatives
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D239/00—Heterocyclic compounds containing 1,3-diazine or hydrogenated 1,3-diazine rings
- C07D239/02—Heterocyclic compounds containing 1,3-diazine or hydrogenated 1,3-diazine rings not condensed with other rings
- C07D239/24—Heterocyclic compounds containing 1,3-diazine or hydrogenated 1,3-diazine rings not condensed with other rings having three or more double bonds between ring members or between ring members and non-ring members
- C07D239/28—Heterocyclic compounds containing 1,3-diazine or hydrogenated 1,3-diazine rings not condensed with other rings having three or more double bonds between ring members or between ring members and non-ring members with hetero atoms or with carbon atoms having three bonds to hetero atoms with at the most one bond to halogen, directly attached to ring carbon atoms
- C07D239/32—One oxygen, sulfur or nitrogen atom
- C07D239/42—One nitrogen atom
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D277/00—Heterocyclic compounds containing 1,3-thiazole or hydrogenated 1,3-thiazole rings
- C07D277/02—Heterocyclic compounds containing 1,3-thiazole or hydrogenated 1,3-thiazole rings not condensed with other rings
- C07D277/20—Heterocyclic compounds containing 1,3-thiazole or hydrogenated 1,3-thiazole rings not condensed with other rings having two or three double bonds between ring members or between ring members and non-ring members
- C07D277/32—Heterocyclic compounds containing 1,3-thiazole or hydrogenated 1,3-thiazole rings not condensed with other rings having two or three double bonds between ring members or between ring members and non-ring members with hetero atoms or with carbon atoms having three bonds to hetero atoms with at the most one bond to halogen, e.g. ester or nitrile radicals, directly attached to ring carbon atoms
- C07D277/38—Nitrogen atoms
- C07D277/42—Amino or imino radicals substituted by hydrocarbon or substituted hydrocarbon radicals
Definitions
- the present invention belongs to the technical field of therapeutic agents, and particularly relates to a therapeutic agent for oral administration or preventive agent for oral administration of pollakiuria and urinary incontinence or a sleep inducer for oral administration using a troborone derivative having excellent oral absorbability.
- Anticholinergics, smooth muscle relaxants and the like have been used as drug treatments for such urinary frequency or urinary incontinence, but smooth muscle relaxants have a weaker action than anticholinergic agents.
- anticholinergic agents develop side effects such as blood pressure and urinary retention.
- age-related changes in human bladder smooth muscle are accompanied by increased atopin pin-resistant contraction. Therefore, anticholinergic agents are insufficient in the effects of urinary incontinence in the elderly.
- WO 99/00366 discloses that trobon derivatives can be used as agents for treating or preventing pollakiuria or urinary incontinence.
- 2-benzyloxy-14-isopropyl-17- [4-phenylpiperazinomethyl] -12,4,6-cycloheptatrien-1-one described in the publication is effective by intravenous injection.
- the publication did not specifically disclose a therapeutic or preventive agent for pollakiuria or urinary incontinence by oral administration.
- the oral route of administration of the drug is oral.
- oral Urinary frequency for administration Antiprophylactic or prophylactic agents are eagerly needed.
- one of the objects of the present invention is to provide a therapeutic or prophylactic agent for pollakiuria and urinary incontinence for oral administration having excellent oral absorbability.
- sleep disorders have become a major social problem in modern society.
- drug treatments for such sleep disorders such as insomnia, barbiturates and benzodiazepines, which are sleep-inducing agents, are used clinically.
- the barbituric acid system has drawbacks such as its toxicity, resistance to drugs and formation of dependence, and is not widely used at present.
- benzodiazepines are used in the treatment of many patients because they have a wider therapeutic range and higher safety than valpiturates, but they form a dependence on the drug, interact with alcohol, Since it causes side effects such as memory impairment and muscle relaxation, safer sleep-inducing agents are desired.
- Another object of the present invention is to provide a sleep inducer for oral administration which has excellent oral absorbability and has few side effects.
- the present inventors have conducted intensive studies to achieve the above object, and as a result, the trobolone derivative shown in the present invention has unexpectedly high bioavailability (bioavailability) and is frequently absorbed from the digestive tract.
- the present inventors have found that the present invention has high activity in treating or preventing urinary incontinence or inducing sleep, and completed the present invention.
- R 1 represents a hydrogen atom or a linear or branched alkyl group having 1 to 3 carbon atoms
- R 2 represents a hydrogen atom or an isopropyl group
- Ar represents a heteroatom.
- a therapeutic or prophylactic agent for oral administration of pollakiuria or urinary incontinence comprising a compound represented by the formula: or a pharmacologically acceptable salt thereof as an active ingredient;
- the compound of the general formula (I) is 4-isopropyl-2-methoxy-7- [4- (2-pyrimidyl) piperazinomethyl] -1,2,4,6-cycloheptanyl-trien-11-one or 2-hydroxy_ 4-1- ⁇ fsopropyl-1 7- [4- (2-pyrimidyl) piperazinomethyl] 1,2,4,6-cycloheptatriene-one for oral administration of pollakiuria and urinary incontinence according to (1) above Therapeutic or prophylactic agents;
- the pharmacologically acceptable salt is DL-tartrate or fumarate
- a sleep-inducing agent for oral administration comprising the compound according to any one of the above (1) to (4) or a pharmacologically acceptable salt thereof as an active ingredient;
- a serotonin 1A antagonist comprising, as an active ingredient, the compound according to any one of (1) to (4) or a pharmacologically acceptable salt thereof;
- the compound represented by the above general formula (I) or a pharmacologically acceptable salt thereof used in the present invention exhibits affinity for a receptor for serotonin 1A, which is a kind of brain amine, Since it has serotonin 1A antagonism, it is expected to be effective as a therapeutic or preventive agent for pollakiuria and urinary incontinence or a sleep inducer.
- the therapeutic or preventive agent for oral administration of pollakiuria or urinary incontinence or the sleep inducer for oral administration of the present invention comprises a compound represented by the above general formula (I) or a pharmacologically acceptable salt thereof as an active ingredient.
- R 1 represents a hydrogen atom or a linear or branched alkyl group having 1 to 3 carbon atoms
- R 2 represents a hydrogen atom or an isopropyl group
- r represents an unsubstituted heteroaromatic group having 2 heteroatoms.
- linear or branched alkyl group having 1 to 3 carbon atoms for R 1 include a methyl group, an ethyl group, an n-propyl group, and an isopropyl group.
- R 2 is preferably an isopropyl group.
- hetero atom of the unsubstituted heteroaromatic group having 2 hetero atoms include the same or different two atoms selected from the group consisting of a nitrogen atom, an oxygen atom, a sulfur atom, and the like.
- Preferred heteroaromatic groups include, for example, 3-pyridazinyl group, 2-pyrimidyl group, 2-pyrazinyl group, 2-thiazolyl group, 2-oxoazolyl group, 2-imidazolyl group, and the like.
- a 2-thiazolyl group is particularly preferred.
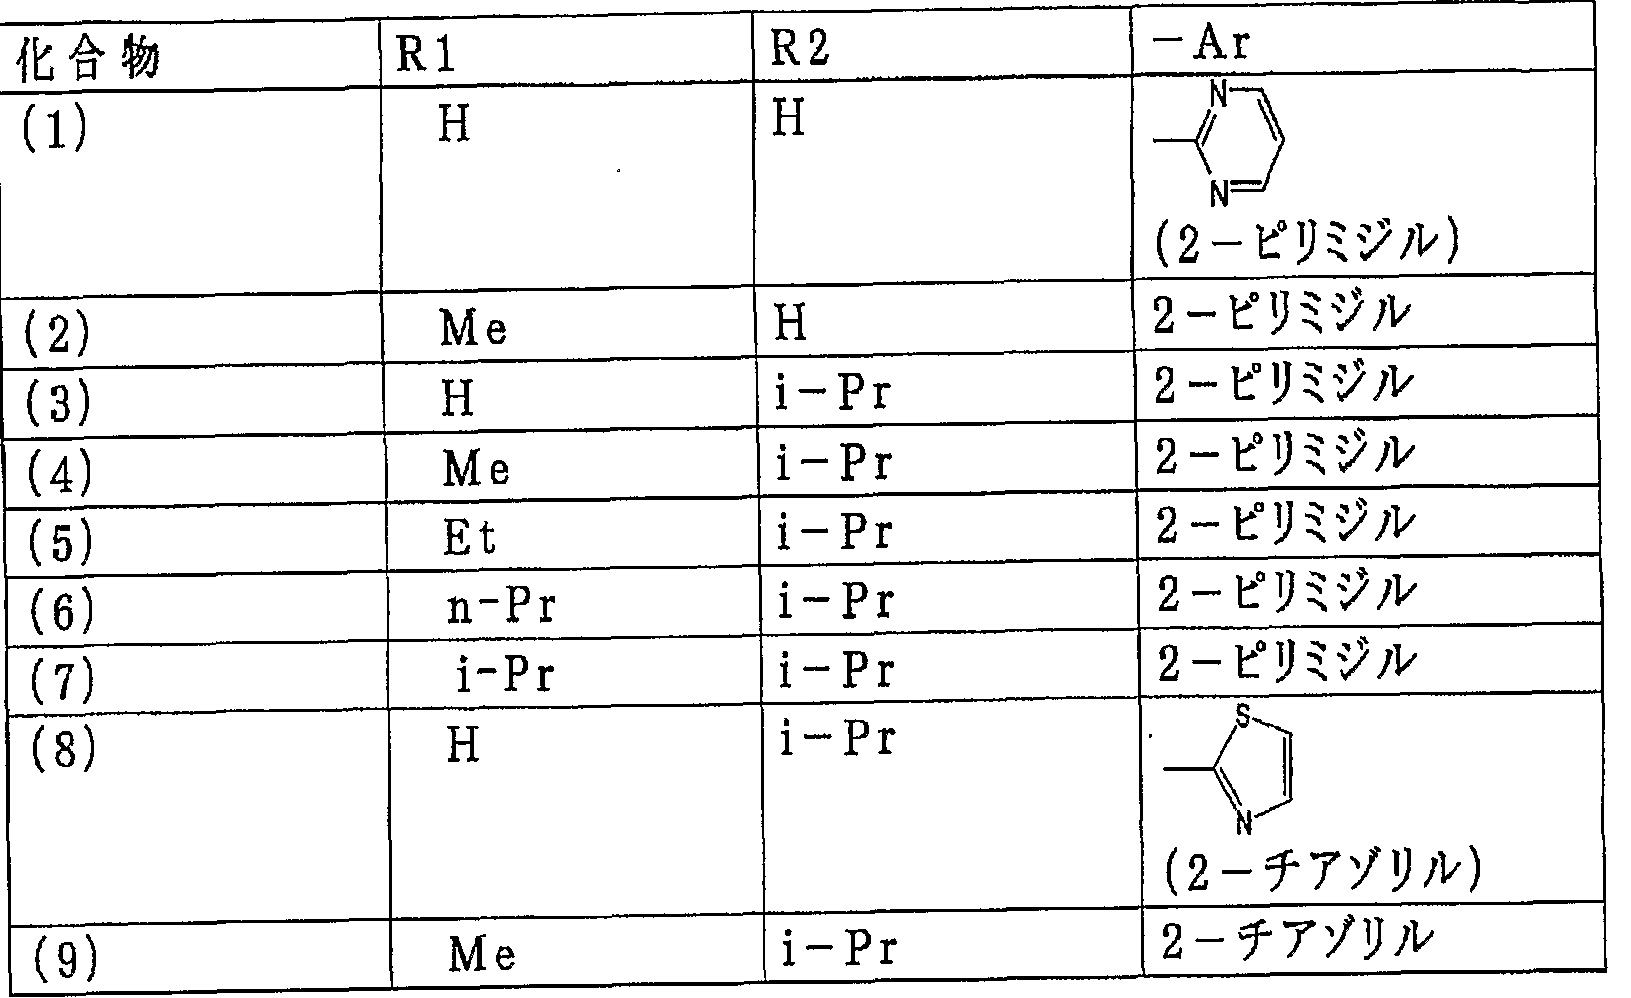
- Table 1 shows examples of the compound represented by the general formula (I).
- H represents a hydrogen atom
- Me represents a methyl group
- Et represents an ethyl group
- n—Pr represents an n-propyl group
- i-Pr represents an isopropyl group.
- the compound of the general formula (I) can be synthesized according to the production method described in WO 99/00366, but is not particularly limited.
- the production route is shown below.
- the amount of pyrimidylpiperazine used is 0.5 to 10 equivalents, preferably 0.8 to 2 equivalents, more preferably 0.8 to 1.2 equivalents to hinokitiol.
- the formalin aqueous solution is used in an amount of 0.5 to 30 equivalents, preferably 0.8 to 2 equivalents.
- the reaction solvent is not particularly limited, but includes alcohols (eg, methanol, ethanol, propanol), esters (eg, methyl acetate, ethyl acetate), ethers (eg, ethyl ether, isopropyl ether, tetrahydrofuran, dioxane), and aromatics.
- Aromatic hydrocarbons (benzene, toluene, xylene, etc.), aliphatic hydrocarbons (pentane, hexane, etc.), water, acetic acid and the like can be mentioned, but alcohols are preferably used.
- the acid include mineral acids such as hydrochloric acid and sulfuric acid, and organic acids such as P-toluenesulfonic acid and acetic acid.
- acetic acid is used.
- the acid is used in an amount of 0.01 equivalent to a solvent amount, preferably 1 to 2 equivalents.
- the reaction temperature is 11 Ot: to around the boiling point of the solvent, preferably from room temperature to around the boiling point of the solvent, and the reaction time is 0.5 to 24 hours, preferably 2 to 8 hours.
- the target compound can be obtained by cooling the reaction solution to precipitate crystals and collecting them by filtration, or by concentrating and recrystallizing, or a general purification method such as column chromatography.
- Methylation of hydroxyl group is performed by adding a base and a methylating agent in an organic solvent.
- a base lithium carbonate, sodium carbonate, potassium carbonate, cesium carbonate, sodium hydroxide, potassium hydroxide, lithium hydroxide, triethylamine, pyridine and the like are used, and the amount used is 0.5 to 10 equivalents, preferably 0 to 10 equivalents. 8 to 2 equivalents.
- the methylating reagent methyl iodide, methyl bromide, dimethyl sulfate and the like are used, and the usage amount is 0.5 to 10 equivalents, preferably 0.8 to 3 equivalents.
- the reaction solvent is not particularly limited, but ketones (acetone, methylethyl ketone, methyl isobutyl ketone, etc.), esters (methyl acetate, ethyl acetate, etc.), ethers (ethyl ether, isopropyl ether, tetrahydrofuran, dioxane, etc.) , Aromatic hydrocarbons (benzene, toluene, xylene, etc.), aliphatic hydrocarbons (pentane, hexane, etc.), and halogenated solvents (methylene salt, methylene chloride, 1,2-dichloroethane, etc.) Dimethylformamide, dimethylsulfoxide and the like.
- ketones acetone, methylethyl ketone, methyl isobutyl ketone, etc.
- esters methyl acetate, ethyl acetate, etc.
- ethers ethyl ether
- the reaction temperature is preferably from 110 ° C. to around the boiling point of the solvent, and the reaction time is preferably from 0.5 to 48 hours.
- a general purification method such as recrystallization or column chromatography. In this way, it is possible to produce 4-isopropyl-12-methoxy-7- [4-1 (2-pyrimidyl) piperazinomethyl] _2,4,6-cycloheptanestrien-1-one (V) .
- the compound (VI) is obtained by reacting hinokitiol with formaldehyde and then performing methylation.
- This is a production method in which the compound is further converted into a compound (VII) by halogenation, and then substituted with pyrimidylpiperazine.
- the pharmacologically acceptable salts of the compound represented by the general formula (I) used in the present invention include salts with mineral acids such as hydrochloric acid and sulfuric acid, methanesulfonic acid, p-toluenesulfonic acid and the like.
- mineral acids such as hydrochloric acid and sulfuric acid, methanesulfonic acid, p-toluenesulfonic acid and the like.
- organic sulfonic acids salts with organic carboxylic acids such as acetic acid, propionic acid, konoic acid, lactic acid, tartaric acid, malic acid, fumaric acid, and maleic acid; with alkali metals such as sodium and potassium And salts with alkaline earth metals such as calcium and magnesium.
- Non-hygroscopic salts such as DL-tartrate or fumarate, specifically 4-Isopropyl-2-methoxy-7- [4-1 (2-pyrimidyl) piperazinomethyl] 1-2,4,6-cyclo Heptatriene-one-one'2 DL-tartrate, 4-isopropyl-2-methoxy-7- [4- (2-pyrimidyl) pyrazinomethyl] -1,2,4,6-cycloheptanyl-one On / 1 fumarate salt is preferred as a drug substance without any problem in guaranteeing its content as an active ingredient.
- a drug having oral absorbability is used for oral administration, and pharmacological activity is obtained by orally administering the drug at a normal dosage that does not show side effects when it has oral absorbability.
- Means to indicate Oral absorbability may be related to absorption from the gastrointestinal tract, transfer into the blood, first-pass effect, distribution to target organs, etc. Effects of intrathecal administration of therapeutic or prophylactic agents or oral sleep-inducing agents on rat rhythmic bladder contraction and the bioavailability of rats.
- the excellent oral absorption of the present invention is shown, and the excellent pharmacological activity effect by oral administration is presumed.
- the therapeutic or preventive agent for urinary frequency and urinary incontinence or the sleep-inducing agent of the present invention comprises the compound alone. It may be used alone or as a composition together with other drugs or medical additives. For example, it can be formulated into oral preparations such as powders, granules, tablets, capsules and the like, if necessary, in combination with pharmaceutically acceptable carriers, excipients, and diluents.
- the content of the compound of the present invention or a pharmaceutically acceptable salt thereof in the preparation is from 0.01 to 100% by weight, preferably from 0.1 to 90% by weight, and the remainder is in the range of a medical additive. .
- the most preferred route of administration is oral (including buccal and sublingual).
- the dosage varies depending on the age of the patient and the condition to be treated. For example, in the case of oral administration to an adult, 0.1 mg to 2000 mg / day, preferably 0.5 mg / day: LO Omg, several times / day It may be administered separately.
- the prophylactic agent for pollakiuria and urinary incontinence means a preparation to be administered before the onset of symptoms in elderly people with frequent urinary frequency and urinary incontinence, and based on slight signs and appropriate indicators. And pharmaceutical preparations to be administered before the onset of symptoms.
- R 1 is a methyl group
- R 2 is an isopropyl group
- Ar is a 2-pyrimidyl group.
- R 1 is a hydrogen atom
- R2 is an isopropyl group
- Ar is a 2-pyrimidyl group 2-hydroxy-4-isopropyl 1 7— [4-1 (2-pyrimidyl) piperazinomethyl] —2,4,6-cycloheptatriene 1-one (compound (3) in Table 1)
- R1 is a hydrogen atom
- R2 is an isopropyl group
- Test Example 1 Test Example 2, Test Example 3, and Test Example 4.
- Test Drug 2 2-Hydroxy-4-Isopropyl-7- [4- (2-Pyrimidyl) piperazinomethyl] -1,2,4,6-cycloheptatrien-1-one / 1-hydrochloride (Compound 'of Example 2; hydrochloride of compound (3) in Table 1)
- Control drug 2_benzyloxy-4-1-isopropyl-17- [4-phenylpyperidinomethyl] -2,4,6-cycloheptatrien-1-one'1 fumarate
- the control drug is described in W099X00366
- Test Example 1 Effect on rat rhythmic bladder contraction
- the anti-micturition effect was evaluated by its effect on rhythmic bladder contraction according to the method of Kaseda et al. (Clinical Physiology, 5, 540-547 (1975)).
- SD-IGS male rats were anesthetized by intraperitoneal administration of urethane (lg / kg). After a midline incision was made in the animal's abdomen to expose the bladder, a small incision was made in the top of the bladder and a rubber balun was inserted therefrom. A polyethylene tube is attached to the balloon, and a three-way stopcock is connected to the other end.A syringe for injecting water to adjust the pressure inside the balloon is used on one side, and a bladder pressure is measured on the other side. A pressure transducer was connected. After leaving the specimen temporarily, water was injected into the balloon until a rhythmic contraction of the bladder was obtained, and the change in the intravesical pressure at this time was continuously recorded in a polygraph. The drug was dissolved in deionized water and administered via a catheter inserted into the duodenum. The effect of the drug was expressed as the inhibition rate (%) of the rhythmic bladder contraction frequency calculated by the following formula.
- Inhibition rate (%) 100- ⁇ Rhythmic contraction frequency for 120 minutes after administration (converted per unit time) / Rhythmic contraction frequency for 20 minutes immediately before administration (converted per unit time) X I 00 ⁇
- test animals three 6-week-old male SD-IGS rats (Nippon Charil Riva Co., Ltd.) were used in each of the intravenous administration group and the oral administration group.
- Each test substance was dissolved in physiological saline or ultrapure water, adjusted to 6.25 mg / kg, and administered intravenously or orally.
- blood is collected over time, plasma is prepared by normal procedures, subjected to protein removal, and the concentration of each test substance is determined using the LCZES I (+) SRM method (liquid chromatography Z I3 ⁇ 4J was determined by spray ionization (+) selected reaction / monitoring method, and the ratio of the area under the blood concentration-time curve at oral administration to the AUC at intravenous administration was expressed as a percentage. Bioavailability was sought.
- Test drugs 1 and 2 had a bioavailability of about 30% to 60% by oral administration, indicating that they were orally available, whereas the control drug had a bioavailability of 6%. Oral administration is unsuitable.
- test drug 1 in a capsule to male 'Beagle' and observation of symptoms did.
- the dose was 3 mgZkg, 1 Omg kg, 3 Omg / kgl O Omg. Kg.
- test drug 1 was prepared using CHO cells transfected with human serotonin 1A receptor. Was tested for its affinity for the human serotonin 1A receptor.
- concentration of test drug 1 was substantially equal to 10- 6 M and plasma concentration at the time of oral administration of 6. 2 5 mgZ kg showing the inhibitory effect of significant rhythmic bladder contractions in rats.
- Test drug of 10- 6 M 1 is the binding to human serotonin 1 A receptor 0. 3 nM of 8- OH @ - DPAT antagonistically inhibits 77%, a strong parent sum Hitose port phosphatonin 1 A receptor Has the property. Subsequently, the results of evaluating the hygroscopicity of the drug of the present invention will be shown.
- Sample 1 (compound of Example 5): 4-Isopropyl-1-2-methoxy_7- [4- (2-pyrimidyl) piperazinomethyl] —2,4,6-cycloheptatrien-1-one′2DL—tartrate
- Sample 2 (Compound of Example 6): 4-Isopropyl-12-methoxy-7- [4- (2-Pyrimidyl) piperazinomethyl] 2,4,6-cycloheptatriene-one-one-one-fumarate
- the present invention will be specifically described by way of examples, but the present invention is not limited thereto.
- the NMR values in the examples are ⁇ values (ppm) measured using tetramethylsilane as an internal standard.
- Example 3 4-Isopropyl-1-2-methoxy-7- [4- (2-pyrimidyl) piperazinomethyl] —2,4,6-cycloheptutrien-1-one obtained in Example 3 (1.0 g, 2.8 mm o 1) was dissolved in 15 mL of ethanol, and an ethanol solution (10 mL) of fumaric acid (0.33 g, 2.8 mmo 1) was added dropwise. After concentration under reduced pressure, 1.5 mL of water and 3 mL of acetone were added and dissolved by heating.
- Hinokitiol (0.29 g, 1.8 mmol), 1— (2-thiazolyl) piperazine (0.30 g, 1.8 mmo1), Acetic acid (0.1 mL, 1,8 mm ⁇ 1) ) was added with 1 OmL of methanol, and a 37% aqueous solution of formalin (0.16 mL, 1.8 mmol 1) was further added. After stirring overnight at room temperature, the precipitated crystals were collected by filtration to give 2-hydroxy-4-isopropyl-17- [4- (2-thiazolyl) piradinomethyl] -2,4,6-cycloheptatriene-1 One ounce (0.3 g, 0.87 mmo 1, 49%) was obtained.
- the anticholinergic agent used as a therapeutic agent for pollakiuria and urinary incontinence does not exhibit side effects such as urinary retention and blood loss, has good oral absorbability, and has high bioavailability.
- Potent frequent urinary frequency urinary incontinence therapeutic or prophylactic agent with minimal metabolic variability among individuals and high response rate expected, especially excellent in vivo stability with few side effects 2-Methoxy-1- 7- [4- (2-Pyrimidyl) piperazinomethyl] -2,4,6-cycloheptanyl-trien-1-one or 2-hydroxy- 4-isopropyl-1 7- [4- (2- Pyrimidyl) piperazinomethyl] -2,4,6-cycloheptatriene-1one or 2-hydroxy-4-1-isopropyl-17- [4- (2-thiazolyl) piperazinomethyl] -1,2,4,6-cycloheptatriene-11one
- the compound causes somnolence in mammals by oral administration, but has no affinity for benzodiazepine receptor and GABA receptor, and has no muscle-relaxing action.
- a short-term sleep-inducing agent that does not cause side effects of a benzodiazepine drug used as an agent.
Landscapes
- Health & Medical Sciences (AREA)
- Chemical & Material Sciences (AREA)
- Organic Chemistry (AREA)
- Veterinary Medicine (AREA)
- Public Health (AREA)
- General Health & Medical Sciences (AREA)
- Animal Behavior & Ethology (AREA)
- Life Sciences & Earth Sciences (AREA)
- Pharmacology & Pharmacy (AREA)
- Medicinal Chemistry (AREA)
- Epidemiology (AREA)
- Nuclear Medicine, Radiotherapy & Molecular Imaging (AREA)
- General Chemical & Material Sciences (AREA)
- Chemical Kinetics & Catalysis (AREA)
- Bioinformatics & Cheminformatics (AREA)
- Engineering & Computer Science (AREA)
- Urology & Nephrology (AREA)
- Neurosurgery (AREA)
- Neurology (AREA)
- Biomedical Technology (AREA)
- Anesthesiology (AREA)
- Pharmaceuticals Containing Other Organic And Inorganic Compounds (AREA)
Description



Claims

Priority Applications (6)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| AU2003234916A AU2003234916A1 (en) | 2002-05-15 | 2003-05-14 | Oral therapeutic or preventive drugs for pollakiuria and urinary incontinence or oral sleep inducers, containing tropolone derivatives |
| CA002485869A CA2485869A1 (en) | 2002-05-15 | 2003-05-14 | Oral therapeutic or preventive drugs for pollakiuria and urinary incontinence or oral sleep inducers, containing tropolone derivatives |
| JP2004505059A JPWO2003097060A1 (ja) | 2002-05-15 | 2003-05-14 | トロポロン誘導体を用いた経口投与用頻尿・尿失禁の治療剤又は予防剤若しくは経口投与用睡眠導入剤 |
| KR10-2004-7016515A KR20040106345A (ko) | 2002-05-15 | 2003-05-14 | 트로폴론 유도체를 사용한 경구투여용 빈뇨ㆍ요실금의치료제 또는 예방제 혹은 경구투여용 수면도입제 |
| EP03752896A EP1504759A1 (en) | 2002-05-15 | 2003-05-14 | Oral therapeutic or preventive drugs for pollakiuria and urinary incontinence or oral sleep inducers, containing tropolone derivatives |
| US10/514,737 US20050130982A1 (en) | 2002-05-15 | 2003-05-14 | Oral therapeutic or preventative drugs for pollakiuria and urinary incontinence and oral sleep inducers, containing tropolone derivatives |
Applications Claiming Priority (4)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| JP2002/140500 | 2002-05-15 | ||
| JP2002140500 | 2002-05-15 | ||
| JP2002/378797 | 2002-12-27 | ||
| JP2002378797 | 2002-12-27 |
Publications (1)
| Publication Number | Publication Date |
|---|---|
| WO2003097060A1 true WO2003097060A1 (en) | 2003-11-27 |
Family
ID=29552282
Family Applications (1)
| Application Number | Title | Priority Date | Filing Date |
|---|---|---|---|
| PCT/JP2003/006003 Ceased WO2003097060A1 (en) | 2002-05-15 | 2003-05-14 | Oral therapeutic or preventive drugs for pollakiuria and urinary incontinence or oral sleep inducers, containing tropolone derivatives |
Country Status (7)
| Country | Link |
|---|---|
| US (1) | US20050130982A1 (ja) |
| EP (1) | EP1504759A1 (ja) |
| JP (1) | JPWO2003097060A1 (ja) |
| KR (1) | KR20040106345A (ja) |
| AU (1) | AU2003234916A1 (ja) |
| CA (1) | CA2485869A1 (ja) |
| WO (1) | WO2003097060A1 (ja) |
Families Citing this family (1)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| WO2013192554A1 (en) | 2012-06-22 | 2013-12-27 | University Of Connecticut | Substituted tropolone derivatives and methods of use |
Citations (1)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| WO1999000366A1 (en) * | 1997-06-27 | 1999-01-07 | Nippon Kayaku Kabushiki Kaisha | Remedies/preventives for frequent urination/urinary incontinence and tropone derivatives |
-
2003
- 2003-05-14 US US10/514,737 patent/US20050130982A1/en not_active Abandoned
- 2003-05-14 WO PCT/JP2003/006003 patent/WO2003097060A1/ja not_active Ceased
- 2003-05-14 JP JP2004505059A patent/JPWO2003097060A1/ja active Pending
- 2003-05-14 KR KR10-2004-7016515A patent/KR20040106345A/ko not_active Withdrawn
- 2003-05-14 AU AU2003234916A patent/AU2003234916A1/en not_active Withdrawn
- 2003-05-14 EP EP03752896A patent/EP1504759A1/en not_active Withdrawn
- 2003-05-14 CA CA002485869A patent/CA2485869A1/en not_active Abandoned
Patent Citations (1)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| WO1999000366A1 (en) * | 1997-06-27 | 1999-01-07 | Nippon Kayaku Kabushiki Kaisha | Remedies/preventives for frequent urination/urinary incontinence and tropone derivatives |
Also Published As
| Publication number | Publication date |
|---|---|
| KR20040106345A (ko) | 2004-12-17 |
| EP1504759A1 (en) | 2005-02-09 |
| AU2003234916A1 (en) | 2003-12-02 |
| CA2485869A1 (en) | 2003-11-27 |
| JPWO2003097060A1 (ja) | 2005-09-15 |
| US20050130982A1 (en) | 2005-06-16 |
Similar Documents
| Publication | Publication Date | Title |
|---|---|---|
| CN100406011C (zh) | 含乙酸酰基苯胺衍生物作为活性成分的膀胱活动过度的治疗药物 | |
| PT1877390E (pt) | Compostos benzisoxazole-piridina e métodos para a sua utilização | |
| WO1999020620A1 (en) | Isoquinoline derivative and drug | |
| JPWO1999020620A1 (ja) | イソキノリン誘導体及び医薬 | |
| CN118948849A (zh) | 用于预防和治疗与抗肥胖作用有关的疾病的药物 | |
| JP2012517456A (ja) | 新規なシタグリプチンの塩 | |
| CN102149683A (zh) | 1-(取代苄基)-5-三氟甲基-2-(1h)吡啶酮化合物及其盐,其制备方法及其用途 | |
| JP3157924B2 (ja) | 治療剤 | |
| CN118598820B (zh) | 一种二苯基三嗪类化合物及其制备方法和应用 | |
| TW202527937A (zh) | 多環化合物及其用途 | |
| WO2010135972A1 (zh) | 1-(取代芳基)-5-三氟甲基-2-(1h)吡啶酮化合物及其盐的制备方法及其用途 | |
| CN111943943A (zh) | 3-芳氧基-3-五元杂芳基-丙胺类化合物及其晶型和用途 | |
| WO2020228789A1 (zh) | 3-芳氧基-3-五元杂芳基-丙胺类化合物及其晶型和用途 | |
| TW490465B (en) | Enterokinetic benzamide, the preparation process and the pharmaceutical compositions thereof | |
| CN115304590B (zh) | 2h-苯并三氮唑衍生物及其制备方法及含有它们的药物组合物 | |
| CN115304593B (zh) | 苯并异噻唑化合物及其药物组合物和应用 | |
| TWI225488B (en) | Derivatives of homopiperidinyl substituted benzimidazole analogues | |
| WO2021160134A1 (zh) | 3-芳氧基-3-五元杂芳基-丙胺类化合物的制备方法及晶型 | |
| JP2017031088A (ja) | インドール誘導体及びその医薬用途 | |
| ES2605466T3 (es) | Compuesto de benzacepina | |
| TW200940543A (en) | Maleate, besylate and L-malate salts of 6-(5-chloro-2-pyridyl)-5-[(4-methyl-1-piperazinyl)carbonyloxy]-7-oxo-6,7-dihydro-5H-pyrrolo[3,4-b]pyrazine | |
| WO2021149767A1 (ja) | ヘテロ環誘導体 | |
| US20250333371A1 (en) | Compounds | |
| WO2006121104A1 (ja) | ピペリジン環を有するインドール誘導体の結晶およびその製法 | |
| JPWO2003097060A1 (ja) | トロポロン誘導体を用いた経口投与用頻尿・尿失禁の治療剤又は予防剤若しくは経口投与用睡眠導入剤 |
Legal Events
| Date | Code | Title | Description |
|---|---|---|---|
| AK | Designated states |
Kind code of ref document: A1 Designated state(s): AE AG AL AM AT AU AZ BA BB BG BR BY BZ CA CH CN CO CR CU CZ DE DK DM DZ EC EE ES FI GB GD GE GH GM HR HU ID IL IN IS JP KE KG KP KR KZ LC LK LR LS LT LU LV MA MD MG MK MN MW MX MZ NI NO NZ OM PH PL PT RO RU SC SD SE SG SK SL TJ TM TN TR TT TZ UA UG US UZ VC VN YU ZA ZM ZW |
|
| AL | Designated countries for regional patents |
Kind code of ref document: A1 Designated state(s): GH GM KE LS MW MZ SD SL SZ TZ UG ZM ZW AM AZ BY KG KZ MD RU TJ TM AT BE BG CH CY CZ DE DK EE ES FI FR GB GR HU IE IT LU MC NL PT RO SE SI SK TR BF BJ CF CG CI CM GA GN GQ GW ML MR NE SN TD TG |
|
| 121 | Ep: the epo has been informed by wipo that ep was designated in this application | ||
| DFPE | Request for preliminary examination filed prior to expiration of 19th month from priority date (pct application filed before 20040101) | ||
| WWE | Wipo information: entry into national phase |
Ref document number: 1020047016515 Country of ref document: KR |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 2003752896 Country of ref document: EP Ref document number: 2004505059 Country of ref document: JP |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 2485869 Country of ref document: CA |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 10514737 Country of ref document: US |
|
| WWP | Wipo information: published in national office |
Ref document number: 1020047016515 Country of ref document: KR |
|
| WWP | Wipo information: published in national office |
Ref document number: 2003752896 Country of ref document: EP |
|
| WWW | Wipo information: withdrawn in national office |
Ref document number: 2003752896 Country of ref document: EP |