WO2004101548A1 - Indazole having analgesic activity , - Google Patents

Indazole having analgesic activity , Download PDF

Info

Publication number
WO2004101548A1
WO2004101548A1 PCT/EP2004/004390 EP2004004390W WO2004101548A1 WO 2004101548 A1 WO2004101548 A1 WO 2004101548A1 EP 2004004390 W EP2004004390 W EP 2004004390W WO 2004101548 A1 WO2004101548 A1 WO 2004101548A1
Authority
WO
WIPO (PCT)
Prior art keywords
alkyl
amino
formula
methyl
branched
Prior art date
Legal status (The legal status is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the status listed.)
Ceased
Application number
PCT/EP2004/004390
Other languages
French (fr)
Inventor
Maria Alessandra Alisi
Nicola Cazzolla
Guido Furlotti
Angelo Guglielmotti
Lorenzo Polenzani
Current Assignee (The listed assignees may be inaccurate. Google has not performed a legal analysis and makes no representation or warranty as to the accuracy of the list.)
Angelini Acraf SpA
Original Assignee
Aziende Chimiche Riunite Angelini Francesco ACRAF SpA
Priority date (The priority date is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the date listed.)
Filing date
Publication date
Priority to PL04729106T priority Critical patent/PL1622892T3/en
Priority to KR1020057021768A priority patent/KR101093048B1/en
Priority to US10/549,930 priority patent/US7662836B2/en
Priority to CA2519733A priority patent/CA2519733C/en
Priority to AT04729106T priority patent/ATE501134T1/en
Priority to AU2004238449A priority patent/AU2004238449B2/en
Priority to UAA200509350A priority patent/UA80605C2/en
Priority to EA200501818A priority patent/EA008499B1/en
Priority to HK06107998.1A priority patent/HK1087705B/en
Priority to SI200431647T priority patent/SI1622892T1/en
Priority to JP2006529710A priority patent/JP4865559B2/en
Priority to MXPA05012297A priority patent/MXPA05012297A/en
Application filed by Aziende Chimiche Riunite Angelini Francesco ACRAF SpA filed Critical Aziende Chimiche Riunite Angelini Francesco ACRAF SpA
Priority to EP04729106A priority patent/EP1622892B1/en
Priority to DK04729106.7T priority patent/DK1622892T3/en
Priority to DE602004031726T priority patent/DE602004031726D1/en
Publication of WO2004101548A1 publication Critical patent/WO2004101548A1/en
Priority to IL170769A priority patent/IL170769A/en
Anticipated expiration legal-status Critical
Priority to US12/638,470 priority patent/US8415476B2/en
Ceased legal-status Critical Current

Links

Classifications

    • CCHEMISTRY; METALLURGY
    • C07ORGANIC CHEMISTRY
    • C07DHETEROCYCLIC COMPOUNDS
    • C07D401/00Heterocyclic compounds containing two or more hetero rings, having nitrogen atoms as the only ring hetero atoms, at least one ring being a six-membered ring with only one nitrogen atom
    • C07D401/02Heterocyclic compounds containing two or more hetero rings, having nitrogen atoms as the only ring hetero atoms, at least one ring being a six-membered ring with only one nitrogen atom containing two hetero rings
    • C07D401/12Heterocyclic compounds containing two or more hetero rings, having nitrogen atoms as the only ring hetero atoms, at least one ring being a six-membered ring with only one nitrogen atom containing two hetero rings linked by a chain containing hetero atoms as chain links
    • CCHEMISTRY; METALLURGY
    • C07ORGANIC CHEMISTRY
    • C07DHETEROCYCLIC COMPOUNDS
    • C07D231/00Heterocyclic compounds containing 1,2-diazole or hydrogenated 1,2-diazole rings
    • C07D231/54Heterocyclic compounds containing 1,2-diazole or hydrogenated 1,2-diazole rings condensed with carbocyclic rings or ring systems
    • C07D231/56Benzopyrazoles; Hydrogenated benzopyrazoles
    • AHUMAN NECESSITIES
    • A61MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61KPREPARATIONS FOR MEDICAL, DENTAL OR TOILETRY PURPOSES
    • A61K31/00Medicinal preparations containing organic active ingredients
    • A61K31/33Heterocyclic compounds
    • A61K31/395Heterocyclic compounds having nitrogen as a ring hetero atom, e.g. guanethidine or rifamycins
    • A61K31/435Heterocyclic compounds having nitrogen as a ring hetero atom, e.g. guanethidine or rifamycins having six-membered rings with one nitrogen as the only ring hetero atom
    • A61K31/44Non condensed pyridines; Hydrogenated derivatives thereof
    • A61K31/445Non condensed piperidines, e.g. piperocaine
    • A61K31/4523Non condensed piperidines, e.g. piperocaine containing further heterocyclic ring systems
    • A61K31/454Non condensed piperidines, e.g. piperocaine containing further heterocyclic ring systems containing a five-membered ring with nitrogen as a ring hetero atom, e.g. pimozide, domperidone
    • AHUMAN NECESSITIES
    • A61MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61PSPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
    • A61P25/00Drugs for disorders of the nervous system
    • A61P25/02Drugs for disorders of the nervous system for peripheral neuropathies
    • AHUMAN NECESSITIES
    • A61MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61PSPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
    • A61P25/00Drugs for disorders of the nervous system
    • A61P25/04Centrally acting analgesics, e.g. opioids
    • AHUMAN NECESSITIES
    • A61MEDICAL OR VETERINARY SCIENCE; HYGIENE
    • A61PSPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
    • A61P29/00Non-central analgesic, antipyretic or antiinflammatory agents, e.g. antirheumatic agents; Non-steroidal antiinflammatory drugs [NSAID]

Definitions

  • the present invention relates to an indazole having analgesic activity, a method for the preparation thereof and a pharmaceutical composition containing the same.
  • Chronic pain is very widespread. On average about 20% of the adult population suffers from it, and it is generally associated with clinical conditions characterized by chronic and/or degenerative lesions.
  • Typical examples of pathologies characterized by chronic pain are rheumatoid arthritis, osteoarthritis, fibromyalgia, neuropathies, etc. [Ashburn MA, Pope PS. Management of chronic pain. Lancet 1999; 353: 1865-69]. Chronic pain is often debilitating and is the cause of loss of the capacity for work and poor quality of life. Therefore it also has adverse economic and social consequences.
  • the analgesic drugs currently used in the treatment of chronic pain belong basically to two classes: the non-steroidal anti-inflammatory drugs (NSAIDs), which combine analgesic activity and anti- inflammatory activity, and the opioid analgesics. These classes constitute the basis for the three-step "analgesic scale" suggested by the World Health Organization for drug treatment of pain [Textbook of Pain. 4th edition. PD Wall and R Melzack Eds. Churchill Livingstone, 1999].
  • NSAIDs non-steroidal anti-inflammatory drugs
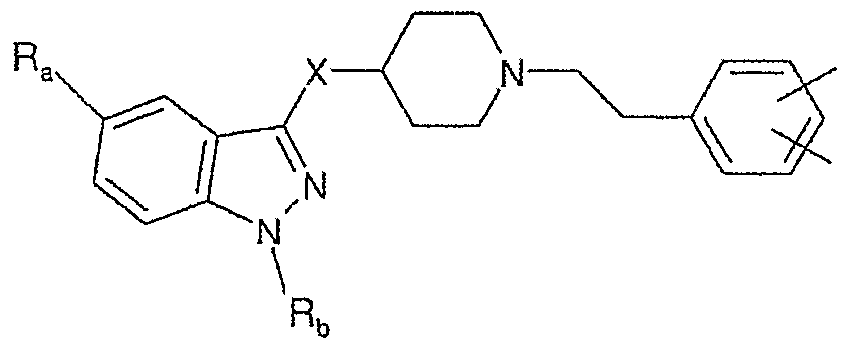
  • the present invention thus relates to an indazole of general formula:
  • X is C(0)NHCH 2 , NHC(O) or NHC(0)CH 2 ;
  • R a is H, NH 2 C(0), CH 3 C(0)NH, CH 3 S0 2 , CH 3 S0 2 NH, linear or branched C C 3 alkyl, linear or branched C 1 -C 3 alkoxy, or halogen;
  • R b is H, linear or branched C- ⁇ -C 6 alkyl; aryl-(C ⁇ -C 3 )alkyI optionally substituted with 1 or 2 halogen atoms, with a C 1 -C 3 alkyl group or a C 1 -C 3 alkoxy group; and in which a) when X is C(0)NHCH 2
  • R G is hydroxy, amino, di-(CrC 3 )alkyl-amino, tri-(CrC- 3 )alkyl- ammoniomethyl, nitro, trifluoromethyl, nitrile, CH 3 C(0)NH, CH3SO2NH, CH3SO2, R'R"NS0 2 , where R' and R" are H, or a linear or branched C C ⁇ alkyl ,
  • R d is H, hydroxy, amino, di-(CrC 3 )alkyl-amino, tri-(CrC 3 )alkyl- ammoniomethyl, nitro, trifluoromethyl, nitrile, CH 3 C(0)NH, CH 3 SO 2 NH, CH3SO2, R'R"NS0 2 , where R 1 and R" have the meanings stated above, with the proviso, however, that when R a and R are both H, and R b is isopropyl, then R c is not hydroxy; b) when X is NHC(O) or NHC(0)CH 2
  • R c and R d which may be equal or different, are H, hydroxy, C 1 -C 3 alkoxy, halogen, amino, di-(C ⁇ -C 3 )alkylamino, tri-(CrC 3 )alkyl- ammoniomethyl, nitro, trifluoromethyl, nitrile, CH 3 C(0)NH,
  • Typical examples of pharmaceutically acceptable acids are: oxalic, maleic, methanesulphonic, paratoluenesulphonic, succinic, citric, tartaric, lactic, hydrochloric, phosphoric, sulphuric.
  • Preferred meanings of R a are H and C 1 -C 3 alkyl.
  • Preferred meanings of R are H and C 1 -C 3 alkyl.
  • R c are H, N0 2 , NH 2 , OH and C 1 -C- 3 alkoxy.
  • the preferred meaning of R is H.
  • the analgesic activity of the compounds of formula (I) was found by means of two experimental models in the rat: mechanical hyperalgesia induced by CFA and mechanical hyperalgesia in diabetic neuropathy induced by streptozocin.
  • CFA-induced hyperalgesia is a syndrome characterized by the activation of circuits with the task of controlling the inflammatory response and associated with the appearance of conditions that interfere with the perception of pain. Injection of CFA is in fact capable of peripherally inducing the release of specific substances (mediators of the inflammatory response and algogenic agents) responsible for local damage and centrally, at the level of the spinal cord, determining biochemical changes that support the amplification of the perception of pain. As is well known, this model constitutes a valid tool for investigating drugs for use in the treatment of inflammatory pain in man and, in particular, in the control of conditions such as hyperalgesia and allodynia.
  • Typical examples of human pathologies characterized by this type of pain associated with degenerative inflammatory processes are rheumatoid arthritis and osteoarthritis.
  • the diabetic neuropathy induced by streptozocin in the rat represents an insulin-dependent syndrome characterized by a concomitant decrease in the conduction velocity of the motor and sensory nerves and the appearance of a number of anomalies in pain perception.
  • this experimental model constitutes a useful too) for the investigation of drugs for use in the treatment of neuropathic pain in man.
  • the model represents a valid example of a whole host of neuropathic pains characterized by phenomena such as hyperalgesia and allodynia following primary lesions or dysfunctions of the nervous system.
  • Typical examples of human pathologies characterized by dysfunctions of this type and by the presence of neuropathic pain are diabetes, cancer, immunodeficiency diseases, trauma, ischaemias, multiple sclerosis, sciatic neuralgias, neuralgia of the trigeminal nerve and post-herpetic syndromes.
  • R c and Rd have the same meanings as stated above or, when R c or Rd is an amino or alcoholic group, R c and R d may be an amino or alcoholic group protected by a protective group of conventional type, with a derivative of an indazole-carboxylic acid of formula (Ilia)
  • R a and R b have the meanings stated above, and
  • Y is a CI or Br atom, or an OR or OC(0)R group, where R is a linear or branched alkyl having from 1 to 6 carbon atoms, or with a derivative of an indazole-carboxylic acid of formula (I I lb)
  • R a has the meanings stated above, b) cleavage of any possible protective group of the aforesaid amino or alcoholic group, and c) optional formation of an acid addition salt of the indazoleamide of formula (I) with a pharmaceutically acceptable organic or inorganic acid.
  • R a and R b have the meanings stated above, is condensed with a derivative of a carboxylic acid of formula (V)
  • R c and R d have the same meanings as stated above or, when R c or R d is an amino or alcoholic group, R c and R d may be an amino or alcoholic group protected by a protective group of conventional type, and Z is a group C(0)Y or CH 2 C(0)Y in which Y is a CI or Br atom, or an OR or OC(0)R group, where R is a linear or branched alkyl having from 1 to 6 carbon atoms, b') cleavage of any possible protective group of the aforesaid amino or alcoholic group, and c') optional formation of an acid addition salt of the indazoleamide of formula (I) with a pharmaceutically acceptable organic or inorganic acid.
  • R d are an N0 2 group the latter can be reduced to give the corresponding compound of formula (I) in which R c and/or R d are NH 2 .
  • the amine of formula (II) may be obtained according to conventional methods. For example, by alkylation of isonipecotamide with a suitable halide and then reduction of the amide to primary amine (WO 9807728) or by protection of aminomethylpiperidine with benzaldehyde (Synthetic Communications 22(16), 2357-2360, 1992), alkylation with a suitable halide and deprotection.
  • the intermediate of formula (II) in which R c and R d have the meanings stated above is novel. This intermediate therefore is another aspect of the present invention.
  • the compounds of formula (Ilia) and (lllb) may also be obtained according to conventional methods.
  • the compounds of formula (Ilia) in which Y is chlorine may be obtained from the corresponding acid with thionyl chloride (J. Med. Chem., 1976, Vol. 19 (6), pages 778-783)
  • the compounds of formula (Ilia) in which Y is OR or OC(0)R may be obtained by means of known reactions of esterification or of formation of mixed anhydrides (R.C. Larok, Comprehensive Organic Transformations, VCH, pages 965-966).
  • the compounds of formula (lllb) may be obtained according to J.O.C. 1958, Vol. 23 page 621.
  • the compounds of formula (V) may also be obtained according to conventional methods.
  • the compounds of formula (V) in which Y is chlorine may be obtained by saponification of the corresponding esters followed by treatment with thionyl chloride.
  • stages (a) and (a 1 ) are carried out by reacting
  • the reaction temperature is of from 15 to 40 e C.
  • the reaction time is of from 1 to 18 hours.
  • the diluent is aprotic, polar or nonpolar. Even more preferably it is aprotic nonpolar.
  • suitable aprotic nonpolar diluents are the aromatic hydrocarbons, e.g. toluene.
  • suitable aprotic polar diluents are dimethylformamide and dimethylsulphoxide.
  • Suitable organic acceptors of acids are pyridine, triethylamine and the like.
  • suitable inorganic acceptors of acids are alkali carbonates and bicarbonates.
  • cleavage of the protective group of the amino or alcoholic group is preferably carried out by techniques known in the chemistry of protective groups.
  • stages (c) and (c 1 ) are preferably preceded by a stage of isolation of the indazoleamide of formula (I).
  • the present invention relates to a pharmaceutical composition containing an effective amount of a compound of formula (I), or of an addition salt thereof with a pharmaceutically acceptable acid, and at least one pharmaceutically acceptable inert ingredient.
  • a pathologic state that might benefit from treatment with a pharmaceutical composition according to the present invention is chronic pain.
  • this chronic pain is due to chronic lesions or to degenerative processes such as rheumatoid arthritis, osteoarthritis, fibromyalgia, oncologic pain, neuropathic pain and the like.
  • the pharmaceutical compositions of the present invention are prepared in a suitable dosage form.
  • suitable dosage forms are tablets, capsules, coated tablets, granules, solutions and syrups for oral administration; creams, ointments and medicated patches for topic administration; suppositories for rectal administration and sterile solutions for injectable, aerosol or ophthalmic administration.
  • these dosage forms will be formulated in such a way as to provide controlled release over time of the compound of formula (I) or of a salt thereof with a pharmaceutically acceptable acid.
  • the required time of release may be very short, normal or protracted.
  • the dosage forms may also contain other conventional ingredients such as: preservatives, stabilizers, surfactants, buffers, salts for the regulation of the osmotic pressure, emulsifiers, sweetener agents, coloring agents, flavouring agents and the like.
  • the pharmaceutical composition of the present invention may contain other pharmacologically active ingredients whose concomitant is therapeutically useful.
  • the amount of the compound of formula (I) or of the pharmaceutically acceptable acid salt thereof in the pharmaceutical composition of the present invention may vary in a wide range depending on known factors, such as for example the type of disease to be treated, the severity of the disease, the patient's body weight, the dosage form, the chosen route of administration, the number of daily administrations and the efficacy of the chosen compound of formula (i). However, the optimum amount may be easily and routinely determined by a person skilled in the art. Typically, the amount of the compound of formula (I) or of a salt thereof with a pharmaceutically acceptable acid in the pharmaceutical composition of the present invention will be such that it ensures an administration level of from 0.001 to 100 mg/kg/day. Even more preferably, of from 0.1 to 10 mg/kg/day.
  • the dosage forms of the pharmaceutical composition of the present invention can be prepared according to techniques that are well known to the pharmaceutical chemist and comprise mixing, granulation, compression, dissolution, sterilization and the like.
  • Example 1 b 1 -(2-(4-nitrophenyl)ethv ⁇ -4-piperidinylmethane-amine
  • the product of Example 1 a) (8.8 g; 0.044 mol) was dissolved in absolute ethanol (50 ml) and added to a suspension containing 2-(4- nitrophenyl)ethylbromide (10.0 g; 0.044 mol) and anhydrous potassium carbonate (12.1 g; 0.088 mol) in absolute ethanol (100 ml).
  • the suspension thus obtained was boiled under reflux for 16 hours.
  • the reaction mixture was then left to cool to room temperature and filtered.
  • the filtrate was evaporated at reduced pressure.
  • Example 5b The product of Example 5b) (1.68 g; 0.006 mol) was added to a solution of 1 -methyl-1 H-3-indazoleamine (0.86 g; 0.006 mol), prepared as described in the Journal of Heterocyclic Chemistry 1979 (16), 783- 784, and of triethylamine (2.4 ml; 0.018 mol) in toluene (20 ml).
  • the reaction mixture was stirred at room temperature for 18 h and then the solvent was removed by evaporation at reduced pressure.
  • the residue thus obtained was taken up with 1 N NaOH and dichloromethane and transferred to a separatory funnel.
  • the organic phase was separated, dried over Na 2 S0 4 and the solvent was removed by evaporation at reduced pressure.
  • the product thus obtained was transformed into the corresponding hydrochloride by dissolution in ethanol, addition of hydrogen chloride in ethanol and recrystallization from ethanol, to give the desired salt (1.6 g). m.p.: 235-2
  • Example 1 b The title product was obtained (9.3 g) by working in a similar way to that described in Example 1 b) but starting from the product of Example 1 a) (7.5 g, 0.037 mol) and 2-(4-hydroxyphenyl)ethyl bromide (7.5 g; 0.037 mol), prepared as described in Acta Chemica Scandinava (1947- 1973) 1967, 21 (1 ), 52-62, instead of from 2-(4-nitrophenyl)ethyl bromide.
  • reaction mixture was stirred for 6 hours at 70 9 C. After cooling, water was added.
  • the reaction mixture was transferred to a separatory funnel and extracted with diethyl ether.
  • the organic phase was washed with water saturate with sodium bicarbonate and, finally, the solvent was removed by evaporation at reduced pressure.
  • Example 8d Working in a similar way to that described in Example 3 but using the product of Example 8a) (4.0 g, 0.017 mol) and the chloride of 1 -(1 - methylethyl)-5-methyl-1 H-indazole-3-carboxylic acid (4.0 g, 0.017 mol), prepared as described in the preceding Example 8d), 4.5 g of the desired product were obtained, and this were transformed into the corresponding hydrochloride by dissolution in absolute ethanol, addition of hydrogen chloride in ethanol and recrystallization from ethanol to give the desired salt (3.2 g). m.p.: 257.5-259.5 9 C Elemental analysis for C 26 H 3 N 0 2 HCI
  • Hyperalgesia was induced by unilateral injection of 150 ⁇ l of Complete Freund's Adjuvant (CFA) in the plantar surface of the animal's left hind foot [Andrew D, Greenspan JD. Mechanical and heat sensitization of cutaneous nociceptors after peripheral inflammation in the rat. J Neurophysiol 1999; 82(5): 2649-2656; Hargreaves K, Dubner R, Brown R, Flores C, Joris J. A new and sensitive method for measuring thermal nociception in cutaneous hyperalgesia. Pain 1988; 32: 77-88]. The compounds under examination were tested (dose 10 "5 mol/kg) by carrying out the test 23 hours after injection of CFA.
  • CFA Complete Freund's Adjuvant
  • Diabetic syndrome was induced by a single intraperitoneal (i.p.) injection of 80 mg/kg of streptozotocin dissolved in sterile physiological solution [Courteix C, Eschalier A, Lavarenne J. Streptozotocin-induced diabetic rats: behavioural evidence for a model of chronic pain. Pain, 1993; 53: 81 -88; Bannon AW, Decker MW, Kim Dj, Campbell JE, Arneric SP. ABT-594, a novel cholinergic channel modulator, is efficacious in nerve ligation and diabetic neuropathy models of neuropathic pain. Brain Res. 1998; 801 : 158-63].
  • rats were selected having a level of glycaemia > 300 mg/dl and having a response threshold to a mechanical nociceptive stimulus ⁇ 120 g.
  • the glycaemia levels were measured by means of a reflectometer using reactive strips impregnated with glucose oxidase.
  • the pain threshold was measured using an analgesiometer. By applying a gradual increase in pressure on the dorsal zone of the rat's left hind foot, the instrument makes it possible to record the nocifensive response, expressed in grams, corresponding to the moment when the animal retracts its foot.
  • the pain threshold measured in control animals was compared with that measured in animals treated with the product under examination (dose 10 "5 mol/kg).
  • the pain threshold of normal animals of equal weight/age 240 ⁇ 8.7 g

Landscapes

  • Chemical & Material Sciences (AREA)
  • Health & Medical Sciences (AREA)
  • Organic Chemistry (AREA)
  • Life Sciences & Earth Sciences (AREA)
  • Veterinary Medicine (AREA)
  • Public Health (AREA)
  • General Health & Medical Sciences (AREA)
  • Medicinal Chemistry (AREA)
  • Animal Behavior & Ethology (AREA)
  • Pharmacology & Pharmacy (AREA)
  • General Chemical & Material Sciences (AREA)
  • Nuclear Medicine, Radiotherapy & Molecular Imaging (AREA)
  • Chemical Kinetics & Catalysis (AREA)
  • Pain & Pain Management (AREA)
  • Engineering & Computer Science (AREA)
  • Bioinformatics & Cheminformatics (AREA)
  • Biomedical Technology (AREA)
  • Neurology (AREA)
  • Neurosurgery (AREA)
  • Rheumatology (AREA)
  • Epidemiology (AREA)
  • Pharmaceuticals Containing Other Organic And Inorganic Compounds (AREA)
  • Plural Heterocyclic Compounds (AREA)

Abstract

An indazole having analgesic activity, a method for the preparation thereof and a pharmaceutical composition containing the same; the indazole has the following general formula: where X, Ra, Rb, Rc and Rd have the meanings stated in the description.

Description

INDAZOLES HAVING ANALGESIC ACITIVY
DESCRIPTION The present invention relates to an indazole having analgesic activity, a method for the preparation thereof and a pharmaceutical composition containing the same.
Chronic pain is very widespread. On average about 20% of the adult population suffers from it, and it is generally associated with clinical conditions characterized by chronic and/or degenerative lesions.
Typical examples of pathologies characterized by chronic pain are rheumatoid arthritis, osteoarthritis, fibromyalgia, neuropathies, etc. [Ashburn MA, Staats PS. Management of chronic pain. Lancet 1999; 353: 1865-69]. Chronic pain is often debilitating and is the cause of loss of the capacity for work and poor quality of life. Therefore it also has adverse economic and social consequences.
The analgesic drugs currently used in the treatment of chronic pain belong basically to two classes: the non-steroidal anti-inflammatory drugs (NSAIDs), which combine analgesic activity and anti- inflammatory activity, and the opioid analgesics. These classes constitute the basis for the three-step "analgesic scale" suggested by the World Health Organization for drug treatment of pain [Textbook of Pain. 4th edition. PD Wall and R Melzack Eds. Churchill Livingstone, 1999].
Chronic pain is notoriously difficult to treat using the treatments currently available. Consequently the development of new analgesics has always been one of the main aims of the pharmaceutical industry.
Nevertheless, despite the extensive research efforts directed towards identifying a suitable analgesic compound, there is a significant number of patients whose pain condition is still not being adequately treated [Scholz J, Woolf CJ. Can we conquer pain? Nat Neusci. 2002; 5: 1062- 76].
Surprisingly, it has now been found that such properties are possessed by a new family of indazoles.
In its first aspect, the present invention thus relates to an indazole of general formula:
Figure imgf000003_0001
(l)
where X is C(0)NHCH2, NHC(O) or NHC(0)CH2;
Ra is H, NH2C(0), CH3C(0)NH, CH3S02, CH3S02NH, linear or branched C C3 alkyl, linear or branched C1-C3 alkoxy, or halogen; Rb is H, linear or branched C-ι-C6 alkyl; aryl-(Cι-C3)alkyI optionally substituted with 1 or 2 halogen atoms, with a C1-C3 alkyl group or a C1-C3 alkoxy group; and in which a) when X is C(0)NHCH2
RG is hydroxy, amino, di-(CrC3)alkyl-amino, tri-(CrC-3)alkyl- ammoniomethyl, nitro, trifluoromethyl, nitrile, CH3C(0)NH, CH3SO2NH, CH3SO2, R'R"NS02, where R' and R" are H, or a linear or branched C Cβ alkyl ,
Rd is H, hydroxy, amino, di-(CrC3)alkyl-amino, tri-(CrC3)alkyl- ammoniomethyl, nitro, trifluoromethyl, nitrile, CH3C(0)NH, CH3SO2NH, CH3SO2, R'R"NS02, where R1 and R" have the meanings stated above, with the proviso, however, that when Ra and R are both H, and Rb is isopropyl, then Rc is not hydroxy; b) when X is NHC(O) or NHC(0)CH2
Rc and Rd, which may be equal or different, are H, hydroxy, C1-C3 alkoxy, halogen, amino, di-(Cι-C3)alkylamino, tri-(CrC3)alkyl- ammoniomethyl, nitro, trifluoromethyl, nitrile, CH3C(0)NH,
CH3SO2NH, CH3SO2, R'R"NS02, where R' and R" have the meanings stated above, and their acid addition salts with pharmaceutically acceptable organic and inorganic acids.
Typical examples of pharmaceutically acceptable acids are: oxalic, maleic, methanesulphonic, paratoluenesulphonic, succinic, citric, tartaric, lactic, hydrochloric, phosphoric, sulphuric. Preferred meanings of Ra are H and C1-C3 alkyl. Preferred meanings of R are H and C1-C3 alkyl.
Preferred meanings of Rc are H, N02, NH2, OH and C1-C-3 alkoxy. The preferred meaning of R is H.
The analgesic activity of the compounds of formula (I) was found by means of two experimental models in the rat: mechanical hyperalgesia induced by CFA and mechanical hyperalgesia in diabetic neuropathy induced by streptozocin.
As is known to a person skilled in the art, the aforementioned experimental models can be regarded as predictive of activity in man.
CFA-induced hyperalgesia is a syndrome characterized by the activation of circuits with the task of controlling the inflammatory response and associated with the appearance of conditions that interfere with the perception of pain. Injection of CFA is in fact capable of peripherally inducing the release of specific substances (mediators of the inflammatory response and algogenic agents) responsible for local damage and centrally, at the level of the spinal cord, determining biochemical changes that support the amplification of the perception of pain. As is well known, this model constitutes a valid tool for investigating drugs for use in the treatment of inflammatory pain in man and, in particular, in the control of conditions such as hyperalgesia and allodynia.
Typical examples of human pathologies characterized by this type of pain associated with degenerative inflammatory processes are rheumatoid arthritis and osteoarthritis.
In its turn, the diabetic neuropathy induced by streptozocin in the rat represents an insulin-dependent syndrome characterized by a concomitant decrease in the conduction velocity of the motor and sensory nerves and the appearance of a number of anomalies in pain perception. As is well known, this experimental model constitutes a useful too) for the investigation of drugs for use in the treatment of neuropathic pain in man. In particular, the model represents a valid example of a whole host of neuropathic pains characterized by phenomena such as hyperalgesia and allodynia following primary lesions or dysfunctions of the nervous system. Typical examples of human pathologies characterized by dysfunctions of this type and by the presence of neuropathic pain are diabetes, cancer, immunodeficiency diseases, trauma, ischaemias, multiple sclerosis, sciatic neuralgias, neuralgia of the trigeminal nerve and post-herpetic syndromes.
In a second aspect, the present invention relates to a method of preparation of a compound of formula (I) in which X = C(0)NHCH and acid addition salts thereof with pharmaceutically acceptable organic or inorganic acids, characterized in that it comprises the following stages: a) reaction of an amine of formula (II)
Figure imgf000006_0001
(ID
where
Rc and Rd have the same meanings as stated above or, when Rc or Rd is an amino or alcoholic group, Rc and Rd may be an amino or alcoholic group protected by a protective group of conventional type, with a derivative of an indazole-carboxylic acid of formula (Ilia)
Figure imgf000006_0002
(Ilia)
where
Ra and Rb have the meanings stated above, and
Y is a CI or Br atom, or an OR or OC(0)R group, where R is a linear or branched alkyl having from 1 to 6 carbon atoms, or with a derivative of an indazole-carboxylic acid of formula (I I lb)
Figure imgf000006_0003
where
Ra has the meanings stated above, b) cleavage of any possible protective group of the aforesaid amino or alcoholic group, and c) optional formation of an acid addition salt of the indazoleamide of formula (I) with a pharmaceutically acceptable organic or inorganic acid.
In a third aspect, the present invention relates to a method of preparation of a compound of formula (I) in which X = NH(CO) or
NH(CO)CH2 and acid addition salts thereof with pharmaceutically acceptable organic or inorganic acids, characterized in that it comprises the following stages: a') reaction of an amine of formula (IV)
Figure imgf000007_0001
where
Ra and Rb have the meanings stated above, is condensed with a derivative of a carboxylic acid of formula (V)
Figure imgf000007_0002
(V)
where
Rc and Rd have the same meanings as stated above or, when Rc or Rd is an amino or alcoholic group, Rc and Rd may be an amino or alcoholic group protected by a protective group of conventional type, and Z is a group C(0)Y or CH2C(0)Y in which Y is a CI or Br atom, or an OR or OC(0)R group, where R is a linear or branched alkyl having from 1 to 6 carbon atoms, b') cleavage of any possible protective group of the aforesaid amino or alcoholic group, and c') optional formation of an acid addition salt of the indazoleamide of formula (I) with a pharmaceutically acceptable organic or inorganic acid. A person skilled in the art will easily understand that some compounds of formula (I) can also be prepared from another compound of formula (I) by conventional techniques. For example, when R0 and/or
Rd are an N02 group the latter can be reduced to give the corresponding compound of formula (I) in which Rc and/or Rd are NH2.
The amine of formula (II) may be obtained according to conventional methods. For example, by alkylation of isonipecotamide with a suitable halide and then reduction of the amide to primary amine (WO 9807728) or by protection of aminomethylpiperidine with benzaldehyde (Synthetic Communications 22(16), 2357-2360, 1992), alkylation with a suitable halide and deprotection. The intermediate of formula (II) in which Rc and Rd have the meanings stated above is novel. This intermediate therefore is another aspect of the present invention.
The compounds of formula (Ilia) and (lllb) may also be obtained according to conventional methods. For example, the compounds of formula (Ilia) in which Y is chlorine may be obtained from the corresponding acid with thionyl chloride (J. Med. Chem., 1976, Vol. 19 (6), pages 778-783), whereas the compounds of formula (Ilia) in which Y is OR or OC(0)R may be obtained by means of known reactions of esterification or of formation of mixed anhydrides (R.C. Larok, Comprehensive Organic Transformations, VCH, pages 965-966). In their turn, the compounds of formula (lllb) may be obtained according to J.O.C. 1958, Vol. 23 page 621.
In their turn the compounds of formula (IV) may be obtained according to the conventional methods described in the literature, for example in J. of Heterocyclic Chemistry 1979 (16) 783-784, or in J.A.C.S., 1943 (65) 1804-1806.
The compounds of formula (V) may also be obtained according to conventional methods. For example, the compounds of formula (V) in which Y is chlorine may be obtained by saponification of the corresponding esters followed by treatment with thionyl chloride.
Preferably, stages (a) and (a1) are carried out by reacting
- a compound of formula (II) with a compound of formula (Ilia) in which Y is chlorine, or
- a compound of formula (II) with a compound of formula (lllb), or - a compound of formula (IV) with a compound of formula (V) in which Y is chlorine, in the presence of a suitable diluent and at a temperature of from 0 to 140aC, for a time of from 0.5 to 20 hours.
Preferably, the reaction temperature is of from 15 to 40eC. Advantageously, the reaction time is of from 1 to 18 hours.
Preferably the diluent is aprotic, polar or nonpolar. Even more preferably it is aprotic nonpolar. Examples of suitable aprotic nonpolar diluents are the aromatic hydrocarbons, e.g. toluene. Examples of suitable aprotic polar diluents are dimethylformamide and dimethylsulphoxide.
In the embodiments in which a compound of formula (II) is reacted with a compound of formula (Ilia) in which Y is chlorine, or in which a compound of formula (IV) is reacted with a compound of formula (V) in which Y is chlorine, the aforesaid stages (a) and, respectively, (a1) are preferably carried out in the presence of an organic or inorganic acceptor of acids.
Examples of suitable organic acceptors of acids are pyridine, triethylamine and the like. Examples of suitable inorganic acceptors of acids are alkali carbonates and bicarbonates.
In stages (b) and (b'), cleavage of the protective group of the amino or alcoholic group is preferably carried out by techniques known in the chemistry of protective groups.
In their turn, stages (c) and (c1) are preferably preceded by a stage of isolation of the indazoleamide of formula (I).
In a further aspect, the present invention relates to a pharmaceutical composition containing an effective amount of a compound of formula (I), or of an addition salt thereof with a pharmaceutically acceptable acid, and at least one pharmaceutically acceptable inert ingredient. A typical example of a pathologic state that might benefit from treatment with a pharmaceutical composition according to the present invention is chronic pain. Typically this chronic pain is due to chronic lesions or to degenerative processes such as rheumatoid arthritis, osteoarthritis, fibromyalgia, oncologic pain, neuropathic pain and the like.
Preferably, the pharmaceutical compositions of the present invention are prepared in a suitable dosage form.
Examples of suitable dosage forms are tablets, capsules, coated tablets, granules, solutions and syrups for oral administration; creams, ointments and medicated patches for topic administration; suppositories for rectal administration and sterile solutions for injectable, aerosol or ophthalmic administration.
Advantageously, these dosage forms will be formulated in such a way as to provide controlled release over time of the compound of formula (I) or of a salt thereof with a pharmaceutically acceptable acid. Indeed, depending on the type of treatment, the required time of release may be very short, normal or protracted.
The dosage forms may also contain other conventional ingredients such as: preservatives, stabilizers, surfactants, buffers, salts for the regulation of the osmotic pressure, emulsifiers, sweetener agents, coloring agents, flavouring agents and the like.
Moreover, when required by particular therapies, the pharmaceutical composition of the present invention may contain other pharmacologically active ingredients whose concomitant is therapeutically useful.
The amount of the compound of formula (I) or of the pharmaceutically acceptable acid salt thereof in the pharmaceutical composition of the present invention may vary in a wide range depending on known factors, such as for example the type of disease to be treated, the severity of the disease, the patient's body weight, the dosage form, the chosen route of administration, the number of daily administrations and the efficacy of the chosen compound of formula (i). However, the optimum amount may be easily and routinely determined by a person skilled in the art. Typically, the amount of the compound of formula (I) or of a salt thereof with a pharmaceutically acceptable acid in the pharmaceutical composition of the present invention will be such that it ensures an administration level of from 0.001 to 100 mg/kg/day. Even more preferably, of from 0.1 to 10 mg/kg/day. The dosage forms of the pharmaceutical composition of the present invention can be prepared according to techniques that are well known to the pharmaceutical chemist and comprise mixing, granulation, compression, dissolution, sterilization and the like.
The following examples will illustrate the invention, without limiting it in any way. In the following examples, the substituents on the aromatic ring (Rc and Rd) are indicated with numbering in bold.
EXAMPLE 1 (N((1 -(2-(4-nitrophenyl)ethyl)-4-piperidinvπmethvπ-1 H-indazole-3- carboxamide hvdrochloride (AF3R298)
(I, Ra = Rb = Rd = H; Rc = 4-N02; X = C(0)NHCH2) a) N-hexahvdro-4-pyridinylmethyl-N-phenylmethylidene-amine Benzaldehyde (4.6 g; 0.044 mol) was added dropwise to a solution of
4-aminomethylpiperidine (5.0 g; 0.044 mol) in toluene (20 ml). The solution thus obtained was stirred for 3 h at room temperature. Then the solvent was removed by evaporation at reduced pressure and the residue was taken up twice with toluene to give the desired product that was used as such without further purification. b) 1 -(2-(4-nitrophenyl)ethvπ-4-piperidinylmethane-amine The product of Example 1 a) (8.8 g; 0.044 mol) was dissolved in absolute ethanol (50 ml) and added to a suspension containing 2-(4- nitrophenyl)ethylbromide (10.0 g; 0.044 mol) and anhydrous potassium carbonate (12.1 g; 0.088 mol) in absolute ethanol (100 ml). The suspension thus obtained was boiled under reflux for 16 hours. The reaction mixture was then left to cool to room temperature and filtered. The filtrate was evaporated at reduced pressure. The residue thus obtained was then suspended in 3N HCl (50 ml) and stirred for 3 h at room temperature. The solution was then transferred to a separatory funnel and the acid aqueous phase was washed with ethyl acetate (4 x 50 ml), the aqueous phase was then made alkaline by addition of 6N NaOH and extracted with dichloromethane. The organic phase was dried over NaS04 and the solvent was removed by evaporation at reduced pressure to give the desired product (9 g). 1H-NMR (δ, CDCI3 + D20): 1.43-1.50 (m, 3H); 1.76 (d J=12 Hz, 2H); 2.03 (t, J=12 Hz, 2H); 2.67-2.52 (m, 4H); 2.82-3.06 (m, 4H); 7.39 (d, J=9 Hz, 2H); 8.12 (d, J=9 Hz, 2H); 7.95 (quintet, J=1 Hz, 1 H). c) N((1 -(2-(4-nitrophenyl)ethvO-4-piperidinyl)methvO-1 H-indazole-3- carboxamide hvdrochloride
A solution of the product of Example 1b) (8.4 g; 0.032 mol) in toluene (85 ml) was added, using a dropping funnel, to a suspension containing 7H,14H-indazole(2',3':4,5)pyrazine(1 ,2-b)indazole-7,14,dione (4.6 g; 0.016 mol) prepared as described in J.O.C., 1958, Vol. 23, page 621 , in toluene (60 ml). The reaction mixture was stirred at room temperature for 38 hours and then filtered. The solid was separated and added to a stirred saturate solution of NaHC03 (200 ml) for 2 h. The reaction mixture was filtered and the thus obtained solid product was converted into the corresponding hydrochloride by dissolution in absolute ethanol, addition of hydrogen chloride in ethanol and recrystallization from ethanol, to give the desired product (4.2 g). m.p.: 251 -252.5°C Elemental analysis for C22H25N5Q3. HCI
Figure imgf000013_0001
1H-NMR (δ, DMSO): 1.52-2.11 (m, 5H); 2.85-302 (m, 2H); 3.17-3.64 (m, 8H); 7.19-7.28 (m, 1 H); 7.36-7.46 (m, 1 H); 7.53-7.56 (m, 3H); 8.13-8.26 (m, 3H); 8.55 (t, J=6 Hz, 1 H); 10.82 (s broad, 1 H); 13.70 (s, 1 H).
EXAMPLE 2 N((1 -(2-(4-aminophenyl)ethyl)-4-piperidinyl)methyl)-1 H-indazole-3- carboxamide dihvdrochloride (AF3R02) (I, Ra = Rb = Rd = H, Rc = 4-NH2) X = C(0)NHCH2)
A solution of the product of Example 1c) in the form of base (3 g; 0.007 mol) in 95s ethanol (200 ml) was hydrogenated on 10% Pd-C (0.3 g) at 40 psi for 3 hours. The mixture was then filtered and the filtrate was concentrated at reduced pressure. The thus obtained product was crystallized from ethyl acetate and transformed into the corresponding hydrochloride by dissolution in a mixture of ethyl acetate:ethanol = 9:1 and addition of hydrogen chloride in ethanol to give the desired product
(1 -2 g). m.p.: 271 -273sC (decomp.)
Elemental analysis for C22H2 N50. 2HC1 1/2 H20
Figure imgf000014_0001
1H-NMR (δ, DMSO + D20): 1.45-1.66 (m, 2H); 1.80-2.00 (m, 3H); 2.86- 3.14 (m, 4H); 3.19-3.35 (m, 4H); 3.46-3.80 (m, 2H + HDO); 7.22-7.35 (m, 3H); 7.35-7.49 (m, 3H); 7.64 (d, J=9 Hz; 1 H); 8.17 (d, J=9 Hz, 1 H).
Example 3
N((1 -(2-(4-nitrophenv0ethvD-4-piperidinv0methvD-1 -(1 -methylethvD-1 H- indazole-3-carboxamide oxalate (AF3R306) (I, Ra = Rd = H, Rb = i-C3H7, Rc = 4-N02, X = C(0)NHCH2)
1 -(1 -methylethyl)-1 H-indazole-3-carboxylic acid chloride (2.45 g; 0.011 mol), prepared as described in EP-B1 -0 975 623, was added portionwise to a solution of product 1 b) (3.0 g; 0.011 mol) and of triethylamine (4.6 ml; 0.033 mol) in toluene (50 ml). The mixture was stirred at room temperature for 18 h. The solvent was removed by evaporation at reduced pressure. The residue was taken up with 1 N NaOH and dichloromethane. The mixture was transferred to a separatory funnel. The organic phase was separated and dried over Na2S0 . The solvent was removed by evaporation at reduced pressure and the residue thus obtained was purified by flash chromatography, eluting with ethyl acetate to give the desired product (5.5 g) which was then transformed into the corresponding oxalate by dissolution in ethyl acetate, addition of a stoichiometric amount of oxalic acid and recrystallization from ethyl acetate:ethanol = 9:1 , twice, to give the desired salt (3.5 g). m.p.: 98SC (decomp.)
Elemental analysis for C25H31N503. C2H204 1/2 H20
Figure imgf000015_0001
1H-NMR (δ, DMSO + D20): 1.55 (d, J=7 Hz, 6H); 1.44-1.66 (m, 2H); 1.83-2.02 (m, 3H); 2.98 (t, J=12 Hz, 2H); 3.10-3.40 (m, 6H); 3.55 (d, J=12 Hz, 2H); 5.07 (heptet, J=7 Hz, 1 H); 7.28 (t, J=8 Hz, 1 H); 7.46 (t, J=7 Hz, 1 H); 7.59 (d, J=9 Hz; 2H); 7.79 (d, J=8 Hz; 1 H); 8.11 -8.26 (m, 3H); 8.42 (t, J=6 Hz, 1 H).
EXAMPLE 4
N((1 -(2-(4-aminophenyl)ethyl)-4-piperidinyl)methyl)-1 -(1 -methylethyl)-
1 H-indazole-3-carboxamide dihydrochloride (AF3R294)
(I, Ra = Rd = H, Rb = i-C3H7, Rc = 4-NH2, X = C(0)NHCH2) A solution of the product of Example 3, in the form of base, (2.7 g;
0.006 mol) in 95s ethanol (30 ml) was hydrogenated on 10% Pd-C (0.27 g) at 40 psi for 5 hours. The mixture was then filtered and the filtrate was concentrated at reduced pressure. The product thus obtained was transformed into the corresponding hydrochloride by dissolution in ethyl acetate, addition of hydrogen chloride in ethanol and recrystallization from a mixture of ethyl acetate:ethanol = 8:2, to give the desired product
(1 - g). m.p.: 2782C (decomp.)
Elemental analysis for C25H33N50. 2HCI H20
Figure imgf000015_0002
1H-NMR (δ, DMSO): 1.55 (d, J=7 Hz, 6H); 1.45-2.13 (m, 5H); 2.80-3.64 (m, 10H); 5.08 (heptet, J=7 Hz, 1 H); 7.20-7.49 (m, 6H); 7.79 (d, J=9 Hz; 1 H); 8.18 (d, J=9 Hz, 1 H) 8.39 (t, J=6 Hz, 1 H); 9.15-11.18 (m, 4H)
EXAMPLE 5 N-(1 -methyl-1 H-indazol-3-vD-1 -(2-phenylethyl)piperidine-4-carboxamide hydrochloride (AF3R334) (I, Ra = Rc = Rd = H, Rb = CH3, X = NHC(O)) a) 1 -(2-phenylethyl)-4-piperidine-carboxylic acid hydrochloride
A suspension of 1-(2-phenylethyl)-4-carbetoxypiperidine (12.2 g, 0.047 mol) obtained as described in J. Med. Chem. 1996 (39), 749-756, in 1 N NaOH (100 ml) was heated under reflux for 4 h. After cooling to room temperature, the solution was made acid with 6N HCI to pH 2, concentrated by evaporation at reduced pressure, and the solid thus obtained was filtered and dried in a stove under vacuum to give the desired product (12.1 g).
1H-NMR (δ, DMSO + D20): 1.79-2.19 (m, 4H); 2.43-3.74 (m, 9H); 7.18- 7.41 (m, 5H); b) 1 -(2-phenylethyl)-4-piperidinecarbonyl chloride hydrochloride
A suspension of the product of Example 5a) (2.0 g; 0.007 mol) and of thionyl chloride (0.81 ml; 0.011 mol) in toluene (20 ml) was heated under reflux for 3 h. The solvent was then removed by evaporation at reduced pressure and the residue was taken up with toluene (2 x 20 ml) to give the desired product (2.2 g) that was used as such without further purification. c) N-(1 -methyl-1 H-indazol-3-vD-1 -(2-phenylethv0piperidine-4- carboxamide hydrochloride
The product of Example 5b) (1.68 g; 0.006 mol) was added to a solution of 1 -methyl-1 H-3-indazoleamine (0.86 g; 0.006 mol), prepared as described in the Journal of Heterocyclic Chemistry 1979 (16), 783- 784, and of triethylamine (2.4 ml; 0.018 mol) in toluene (20 ml). The reaction mixture was stirred at room temperature for 18 h and then the solvent was removed by evaporation at reduced pressure. The residue thus obtained was taken up with 1 N NaOH and dichloromethane and transferred to a separatory funnel. The organic phase was separated, dried over Na2S04 and the solvent was removed by evaporation at reduced pressure. The product thus obtained was transformed into the corresponding hydrochloride by dissolution in ethanol, addition of hydrogen chloride in ethanol and recrystallization from ethanol, to give the desired salt (1.6 g). m.p.: 235-237eC
Elemental analysis for C22H26N4O.HCI ΛA H2Q
Figure imgf000017_0001
1H-NMR (δ, DMSO + D20): 1.91 -2.27 (m, 4H) 2.70-3.42 (m, 7H); 3.63- 3.75 (m, 2H); 3.96 (s, 3H); 7.10 (t, J=8 Hz, 1 H); 7.22-7.46 (m; 6H); 7.56 (d, J=8 Hz, 1 H) 7.74 (d, J=8 Hz, 1 H); 10.51 (s, 1 H) EXAMPLE 6
N-(1 -methyl-1 H-indazol-3-yl)-1 -(2-(4-methoxyphenyl)ethyl)piperidine-4- carboxamide hydrochloride (AF3R328) (I, Ra = Rd = H, Rb = CH3, Rc = 4-OCH3, X = NHC(O)) a) 1 -(2-(4-methoxyphenyl)ethyl)-4-piperidine-carboxylic acid hydrochloride
The title product was obtained (15.8 g) by working in a similar way to that described in Example 5a) but starting from 1 -(2-(4- methoxyphenyl)ethyl)-4-carbetoxypiperidine (16.5 g; 0.057 mol), prepared as described in US 6 017 931 , instead of from 1 -(2- phenylethyl)-4-carbetoxypiperidine.
1H-NMR (δ, DMSO): 1.80-2.17 (m, 4H); 2.41 -3.74 (m, 7H); 3.73- (s, 3H); 6.89 (d, J=9 Hz, 2H); 7.19 (d, J=9 Hz, 2H) 11.00 (s broad, 1 H); 12.53 (s broad, 1 H) b) 1 -(2-(4-methoxyphenv0ethv0-4-piperidinecarbonyl chloride hydrochloride
The title product was obtained (14.2 g) starting from the product of Example 6a) (13.8 g; 0.048) and by working in a similar way to that described in example 5b). The product thus obtained was used as such without further purification. c) N-(1 -methyl-1 H-indazol-3-yl)-1 -(4-methoxyphenv0ethyl)piperidine-4- carboxamide hydrochloride
The title product was obtained (9.2 g) starting from the product of Example 6b) (14.2 g; 0.045 mol) and from 1 -methyl-1 H-3- indazoleamine (6.6 g; 0.045 mol) and by working in a way similar to that described in Example 5c). A mixture of ethyl acetate:ethanol 9:1 was used as the crystallization solvent. m.p.: 137-1392C (decomp.) Elemental analysis for C23H28N402.HCl H20
Figure imgf000018_0001
H-NMR (δ, DMSO): 1.95-2.25 (m, 4H) 2.69-3.48 (m, 7H); 3.57-3.70 (m, 2H); 3.74 (s, 3H); 3.96 (s, 3H); 6.92 (d, J=9 Hz, 2H); 7.08 (t, J=9 Hz, 1 H); 7.20 (d, J=9 Hz, 2H); 7.38 (t, J=8 Hz, 1 H); 7.56 (d, J=9 Hz, 1 H); 7.76 (d, J=8 Hz, 1 H); 10.36-11.07 (m, 2H) EXAMPLE 7
N-(1 -methyl-1 H-indazol-3-yl)-1 -2-4-hvdroxyphenyl)ethyl)piperidine-4- carboxamide hydrochloride (AF3R330)
(I, Ra = Rd = H, Rb = CH3, Rc = 4-OH, X = NHC(O))
A solution of the product of Example 6c) (6.7 g; 0.017 mol) in dichloromethane (300 ml) was added dropwise to a solution of BBr3
(8.5 g; 0.034 mol) in dichloromethane (50 ml). The reaction mixture was stirred at room temperature for 6 h. Then water was added carefully and the mixture was made alkaline with 1 N NaOH to pH = 9 and transferred to a separatory funnel. The organic phase was separated, dried over Na2S0 , and the solvent was removed by evaporation at reduced pressure. The residue thus obtained (4.4 g) was purified by flash chromatography using a mixture CHCI3:MeOH = 9:1 as eluent. 3 g of product were thus obtained, and were transformed into the corresponding hydrochloride by dissolution in ethanol, addition of hydrogen chloride in ethanol, evaporation of the solvent and crystallization from a mixture of ethyl acetate:ethanol 9:1 , to give the desired product (2.8 g). m.p.: 249-252sC Elemental analysis for C22H26N402.HCl 2/3 H20
Figure imgf000019_0001
1H-NMR (δ, DMSO): 1.96-2.25 (m, 4H) 2.67-3.47 (m, 7H); 3.63 (d, J=12 Hz, 2H); 3.95 (s, 3H); 6.69-6.80 (m, 2H); 7.00-7.13 (m, 3H); 7.38 (t, J=9 Hz, 1 H); 7.56 (d, J=9 Hz, 1 H); 7.76 (d, J=9 Hz, 1 H); 9.37 (s broad, 1 H); 10.35-10.90 (m, 2H)
EXAMPLE 8 N((1 -(2-(4-hvdroxyphenyl)ethyl)-4-piperidinyl)methv0-5-methyl-1 -(1 - methylethyl)-1 H-indazole-3-carboxamide hydrochloride (AF3R296) (I, Ra = CH3, Rb = i-C3H7, Rc = 4-OH, Rd = H, X = C(0)NHCH2) a) 1 -(2-(4-hvdroxyphenvDethvD-4-piperidinylmethane-amine
The title product was obtained (9.3 g) by working in a similar way to that described in Example 1 b) but starting from the product of Example 1 a) (7.5 g, 0.037 mol) and 2-(4-hydroxyphenyl)ethyl bromide (7.5 g; 0.037 mol), prepared as described in Acta Chemica Scandinava (1947- 1973) 1967, 21 (1 ), 52-62, instead of from 2-(4-nitrophenyl)ethyl bromide. 1H-NMR (δ, CDCI3 + D20): 1.15-1.41 (m, 3H); 1.74 (d, J=9 Hz, 2H); 1.90-2.07 (m, 2H); 2.45-2.61 (m,'4H); 2.65-2.75 (m, 2H); 3.01 (d, J=12 Hz, 2H); 6.75 (d, J=9 Hz, 2H); 7.00 (d, J=9 Hz, 2H). b) isopropyl ester of 1-(1-methylethvO-5-methyl-1 H-indazole-3- carboxylic acid
A 60% suspension of sodium iodide in mineral oil (17.1 g; 0.43 mol) was added to a suspension of 5-methyl-1 H-indazole-3-carboxylic acid (30 g; 0.17 mol) prepared as described in J. Heterocyclic Chem. 1964, Vol. 1 (5) 239-241 , in dimethylformamide (450 ml), and the reaction mixture was heated to 70gC. After 30 minutes, isopropyl bromide (48 ml, 0.51 mol) was added.
The reaction mixture was stirred for 6 hours at 709C. After cooling, water was added. The reaction mixture was transferred to a separatory funnel and extracted with diethyl ether. The organic phase was washed with water saturate with sodium bicarbonate and, finally, the solvent was removed by evaporation at reduced pressure.
In this way 20 g of an oil were obtained, which were purified by flash chromatography, eluting with a mixture hexane:ethyl acetate = 7:3, to give 12 g of the desired product. 1H-NMR (δ, CDCI3): 1.47 (d, J=6 Hz, 6H); 1.64 (d, J=7 Hz, 6H); 2.50 (d, J=1 Hz, 3H); 4.92 (heptet, J=7 Hz, 1 H); 5.39 (heptet, J=6 Hz, 1 H); 7.23 (dd, J=9;1 Hz, 1 H); 7.40 (d, J=9 Hz, 1 H); 7.95 (quintet, J=1 Hz, 1 H). c) 1 -d -methylethvP-5-methyl-1 H-indazole-3-carboxylic acid
A suspension of the product prepared according to Example 8b) (8 g; 0.03 mol) in 1 M NaOH (42 ml) was heated under reflux for 3 hours. It was then poured into water, acidified with 2M HCI and extracted with dichloromethane. After evaporation of the solvent at reduced pressure,
7 g of the desired product were obtained. 1H-NMR (δ, CDCI3): 1.61 (d J=7 Hz, 6H); 2.44 (s, 3H); 4.88 (heptet, J=7 Hz, 1 H); 7.19 (d, J=9 Hz, 1 H); 7.34 (d, J=9 Hz, 1 H); 7.97 (s, 1 H); 9.32 (s broad, 1 H). d) chloride of 1 -(1 -methylethvπ-5-methyl-1 H-indazole-3-carboxylic acid Thionyl chloride (6.78 g; 0.057 mol) was added to a suspension of the product prepared according to Example 8c) (4.01 g; 0.019 mol) in toluene (70 ml), and the reaction mixture was heated under reflux for 2 hours. The solvent was removed by evaporation at reduced pressure and taken up twice with toluene (50 ml x 2) to give the desired product (4.3 g) which was used as such without further purification. e) N((1 -(2-(4-hvdroxyphenvOethvO-4-piperidinyl)methyl)-5-methyl-1 -(1 - methylethyl)-1 H-indazole-3-carboxamide hydrochloride
Working in a similar way to that described in Example 3 but using the product of Example 8a) (4.0 g, 0.017 mol) and the chloride of 1 -(1 - methylethyl)-5-methyl-1 H-indazole-3-carboxylic acid (4.0 g, 0.017 mol), prepared as described in the preceding Example 8d), 4.5 g of the desired product were obtained, and this were transformed into the corresponding hydrochloride by dissolution in absolute ethanol, addition of hydrogen chloride in ethanol and recrystallization from ethanol to give the desired salt (3.2 g). m.p.: 257.5-259.59C Elemental analysis for C26H3 N 02 HCI
Figure imgf000021_0001
1H-NMR (δ, DMSO): 1.53 (d, J=7 Hz, 6H); 1.44-1.76 (m, 3H); 1.87 (d, J=12 Hz, 2H); 2.42 (s, 3H); 2.79-3.45 (m, 8H); 3.54 (d, J=12 Hz, 2H); 5.03 (heptet, J=7 Hz, 1 H); 6.73 (d, J=9 Hz, 2H); 7.05 (d, J=9 Hz, 2H); 7.26 (d,d J=9.2 Hz, 1 H); 7.67 (d, J=9 Hz; 1 H); 7.96 (s, 1 H); 8.30 (t, J=6 Hz, 1 H); 9.35 (s, 1 H); 10.35 (s broad, 1 H). TESTS
1. Mechanical hyperalgesia induced by CFA in the rat
Male CD rats weighing 150-200 g on arrival were used. Using an analgesiometer, rats were selected having a response threshold to a , mechanical nociceptive stimulus of from 150 to 180 g. By applying a gradual increase in pressure on the dorsal zone of the rat's left hind foot, the instrument makes it possible to record the nocifensive response, expressed in grams, corresponding to the moment when the animal retracts its foot [Randall LO and Semite JJ. A method for the measurement of analgesic activity on inflamed tissue. Arch. Int. Pharmacodyn. Ther. 1957; 111 : 409-419].
Hyperalgesia was induced by unilateral injection of 150 μl of Complete Freund's Adjuvant (CFA) in the plantar surface of the animal's left hind foot [Andrew D, Greenspan JD. Mechanical and heat sensitization of cutaneous nociceptors after peripheral inflammation in the rat. J Neurophysiol 1999; 82(5): 2649-2656; Hargreaves K, Dubner R, Brown R, Flores C, Joris J. A new and sensitive method for measuring thermal nociception in cutaneous hyperalgesia. Pain 1988; 32: 77-88]. The compounds under examination were tested (dose 10"5 mol/kg) by carrying out the test 23 hours after injection of CFA.
At 1 h following the treatment, the pain threshold measured in control animals was compared with that measured in animals treated with the product under examination. The control animals were treated with the same vehicle (water) used for administering the products under examination. The results are shown in Table 1.
Table 1 - Effect on CFA
Figure imgf000022_0001
Figure imgf000023_0001
The pain threshold of normal animals of equal weight/age = 155 ± 2.1 g 2. Mechanical hyperalgesia in rats with diabetes induced by streptozotocin
Male CD rats weighing 240-300 g on arrival were used. Diabetic syndrome was induced by a single intraperitoneal (i.p.) injection of 80 mg/kg of streptozotocin dissolved in sterile physiological solution [Courteix C, Eschalier A, Lavarenne J. Streptozotocin-induced diabetic rats: behavioural evidence for a model of chronic pain. Pain, 1993; 53: 81 -88; Bannon AW, Decker MW, Kim Dj, Campbell JE, Arneric SP. ABT-594, a novel cholinergic channel modulator, is efficacious in nerve ligation and diabetic neuropathy models of neuropathic pain. Brain Res. 1998; 801 : 158-63].
After at least three weeks following the injection of streptozotocin, rats were selected having a level of glycaemia > 300 mg/dl and having a response threshold to a mechanical nociceptive stimulus < 120 g. The glycaemia levels were measured by means of a reflectometer using reactive strips impregnated with glucose oxidase. The pain threshold was measured using an analgesiometer. By applying a gradual increase in pressure on the dorsal zone of the rat's left hind foot, the instrument makes it possible to record the nocifensive response, expressed in grams, corresponding to the moment when the animal retracts its foot. At 2 h following the treatment, the pain threshold measured in control animals was compared with that measured in animals treated with the product under examination (dose 10"5 mol/kg).
The control animals were treated with the same vehicle (water) used for administering the products under examination. The results are shown in Table 2. Table 2 - Effect on diabetic neuropathy
Figure imgf000024_0001
The pain threshold of normal animals of equal weight/age = 240 ± 8.7 g

Claims

CLAIMS 1. A compound of formula:
Figure imgf000025_0001
where X is C(0)NHCH2, NHC(O) or NHC(0)CH2;
Ra is H, NH2C(0), CH3C(0)NH, CH3S02, CH3S02NH, linear or branched C1-C3 alkyl, linear or branched CrC3 alkoxy, or halogen; Rb is H, linear or branched d-Cβ alkyl; aryi-(Cι-C3)alkyl optionally substituted with 1 or 2 halogen atoms, with a C-1-C3 alkyl group or a C1-C3 alkoxy group; and in which a) when X is C(0)NHCH2
Rc is hydroxy, amino, di-(CrC3)alkyl-amino, tri-(C1-C3)alkyl- ammoniomethyl, nitro, trifluoromethyl, nitrile, CH3C(0)NH,
CH3S02NH, CH3SO2, R'R"NS02, where R' and R" are H, or a linear or branched C C6 alkyl , Rd is H, hydroxy, amino, di-(CrC3)alkyl-amino, tri-(d-
C3)alkylammoniomethyl, nitro, trifluoromethyl, nitrile, CH3C(0)NH, CH3SO2NH, CHgSOs, R'R"NS02, where R' and
R" have the meanings stated above, with the proviso, however, that when Ra and Rd are both H, and Rb is isopropyl, then Rc is not hydroxy; b) when X is NHC(O) or NHC(0)CH2 Rc and R , which may be equal or different, are H, hydroxy, C1-C3 alkoxy, halogen, amino, di-(C-ι-C3)alkylamino, tri-(Cr C3)alkylammoniomethyl, nitro, trifluoromethyl, nitrile, CH3C(0)NH, CH3SO2NH, CH3S02, R'R"NS02) where R' and R" have the meanings stated above, and their acid addition salts with pharmaceutically acceptable organic and inorganic acids. 2. A compound according to claim 1 , characterized in that Ra is H or C1-C3 alkyl. 3. A compound according to claim 1 or 2, characterized in that Rb is H or C1-C3 alkyl.
4. A compound according to any one of the claims 1 to 3, characterized in that Rc is H, N02, NH2, OH or CrC3 alkoxy.
5. A compound according to any one of the claims 1 to 4, characterized in that Rd is H.
6. An acid addition salt of a compound according to any one of the claims 1 to 5, characterized in that the acid is selected from the group comprising oxalic, maleic, methanesulphonic, paratoluenesulphonic, succinic, citric, tartaric, lactic, hydrochloric, phosphoric and sulphuric acid.
7. N((1 -(2-(4-nitrophenyl)ethyl)-4-piperidinyl)methyl)-1 H-indazole-3- carboxamide and the pharmaceutically acceptable acid addition salts thereof.
8. N((1 -(2-(4-nitrophenyl)ethyl)-4-piperidinyl)methyl)-1 H-indazole-3- carboxamide hydrochloride.
9. N((1 -(2-(4-aminophenyl)ethyl)-4-piperidinyl)methyl)-1 H-indazole-3- carboxamide and the pharmaceutically acceptable acid addition salts thereof.
10. N((1 -(2-(4-aminophenyl)ethyl)-4-piperidinyi)methyl)-1 H-indazole-3- carboxamide dihydrochloride.
11. N((1 -(2-(4-nitrophenyl)ethyl)-4-piperidinyl)methyl)-1 -(1 - methylethyl)-1 H-indazole-3-carboxamide and the pharmaceutically acceptable acid addition salts thereof.
12. N((1 -(2-(4-nitrophenyl)ethyl)-4-piperidinyl)methyl)-1 -(1 - methylethyl)-1 H-indazole-3-carboxamide oxalate.
13. N((1 -(2-(4-aminophenyl)ethyl)-4-piperidinyl)methyl)-1 -(1 - methylethyl)-1 H-indazole-3-carboxamide and the pharmaceutically acceptable acid addition salts thereof.
14. N((1 -(2-(4-aminophenyl)ethyI)-4-piperidinyl)methyl)-1 -(1 - methylethyI)-1 H-indazole-3-carboxamide dihydrochloride.
15. N-(1 -methyl-1 H-indazol-3-yl)-1 -(2-phenylethyl)piperidine-4- carboxamide and the pharmaceutically acceptable acid addition salts thereof.
16. N-(1 -methyl-1 H-indazol-3-yl)-1 -(2-phenylethyI)piperidine-4- carboxamide hydrochloride.
17. N-(1 -methyl-1 H-indazol-3-yl)-1 -(2-(4- methoxyphenyl)ethyl)piperidine-4-carboxamide and the pharmaceutically acceptable acid addition salts thereof.
18. N-(1 -methyl-1 H-indazol-3-yl)-1 -(2-(4- methoxyphenyl)ethyl)piperidine-4-carboxamide hydrochloride.
19. N-(1 -methyl-1 H-indazol-3-yl)-1 -(2-(4- hydroxyphenyl)ethyl)piperidine-4-carboxamide and the pharmaceutically acceptable acid addition salts thereof.
20. N-(1 -methyl-1 H-indazol-3-yl)-1 -(2-(4- hydroxyphenyl)ethyl)piperidine-4-carboxamide hydrochloride.
21. N((1 -(2-(4-hydroxyphenyl)ethyl)-4-piperidinyl)methyl)-5-methyl-1 - (1 -methylethyl)-1 H-indazole-3-carboxamide and the pharmaceutically acceptable acid addition salts thereof.
22. N((1 -(2-(4-hydroxyphenyl)ethyl)-4-piperidinyl)methyl)-5-methyl-1 - (1 -methylethyl)-1 H-indazole-3-carboxamide hydrochloride.
3. A method for preparing a compound of formula (I)
Figure imgf000028_0001
(I) and its acid addition salts with pharmaceutically acceptable organic or inorganic acids, where
X is C(0)NHCH2;
Ra is H, NH2C(0), CH3C(0)NH, CH3S02, CH3S02NH, linear or branched C1-C3 alkyl, linear or branched C C3 alkoxy, or halogen; Rb is H, linear or branched CrC6 alkyl; aryl-(CrC3)alkyl optionally substituted with 1 or 2 halogen atoms, with a C-1-C3 alkyl group or a C-1-C3 alkoxy group; Rc is hydroxy, amino, di-(Cι-C3)alkyl-amino, tri-(Cr C3)alkylammoniomethyl, nitro, trifluoromethyl, nitrile, CH3C(0)NH, CH3SO2NH, CH3SO2, R'R"NS02, where R' and
R" are H, or a linear or branched C-ι-C6 alkyl , Rd is H, hydroxy; amino, di-(CrC3)alkyl-amino, tri-(Cr C3)alkylammoniomethyl, nitro, trifluoromethyl, nitrile, CH3C(0)NH, CH3S02NH, CH3S02, R'R"NS02, where R' and R" have the meanings stated above, with the proviso, however, that when Ra and Rd are both H, and Rb is isopropyl, then Rc is not hydroxy; characterized in that it comprises the following stages: a) reaction of an amine of formula (II)
Figure imgf000029_0001
(ii)
where
Rc and Rd have the same meanings as stated above or, when
Rc or Rd is an amino or alcoholic group, Rc and Rd may be an amino or alcoholic group protected by a conventional protective group, with a derivative of an indazole-carboxylic acid of formula (Ilia)
Figure imgf000029_0002
la)
where
Ra and Rb have the meanings stated above, and
Y is a CI or Br atom, or a group OR or OC(0)R, where R is a linear or branched alkyl having 1 to 6 carbon atoms, or with a derivative of an indazole-carboxylic acid of formula
(lllb)
Figure imgf000029_0003
(lllb) where
Ra has the meanings stated above, b) cleavage of any possible protective group of the aforesaid amino or alcoholic group, and c) optional formation of an acid addition salt of the indazolamide of formula (I) with a pharmaceutically acceptable organic or inorganic acid.
24. A method of preparation a compound of formula (1)
Figure imgf000030_0001
(I) and the pharmaceutically acceptable acid addition salts thereofwith organic or inorganic acids, where
X is NHC(O) or NHC(0)CH2;
Ra is H, NH2C(0), CH3C(0)NH, CH3S02, CH3S02NH, linear or branched C1-C3 alkyl, linear or branched C1-C3 alkoxy, or halogen; Rb is H, linear or branched C C6 alkyl; aryl-(C C3)alkyl optionally substituted with 1 or 2 halogen atoms, with a C1-C3 alkyl group or a C1-C3 alkoxy group; Rc and R , which may be equal or different, are H, hydroxy, C C3 alkoxy, halogen, amino, di-(Cι-C3)alkylamino, tri-(d- C3)alkylammoniomethyl, nitro, trifluoromethyl, nitrile, CH3C(0)NH, CH3SO2NH, CH3S02, R'R"NS02, where R1 and R" are H, or linear or branched C-ι-C6 alkyl, characterized in that it comprises the following stages: a') reaction of an amine of formula (IV)
Figure imgf000031_0001
(IV)
where
Ra and R have the meanings stated above, is condensed with a derivative of a carboxylic acid of formula
(V)
Figure imgf000031_0002
(V)
where
Rc and Rd have the same meanings as stated above or, when
Rc or Rd is an amino or alcoholic group, Rc and Rd may be an amino or alcoholic group protected by a protective group of conventional type, and
Z is a group C(0)Y or CH2C(0)Y in which Y is a CI or Br atom, or an OR or OC(0)R group, where R is a linear or branched alkyl having from 1 to 6 carbon atoms, b') cleavage of any possible protective group of the aforesaid amino or alcoholic group, and c') optional formation of a salt of acid addition of the indazolamide of formula (I) with a pharmaceutically acceptable organic or inorganic acid. 25. A method according to claim 23, characterized in that stage (a) is carried out by reacting a compound of formula (II) with a compound of formula (Ilia) in which Y is chlorine, or with a compound of formula (lllb) in the presence of a suitable diluent and at a temperature of from 0 to 1409C for a time of from 0.5 to 20 hours. 26. A method according to claim 24, characterized in that stage (a1) is carried out by reacting a compound of formula (IV) with a compound of formula (V) in which Y is chlorine in the presence of a suitable diluent and at a temperature of from 0 to 140SC for a time of from 0.5 to 20 hours. 27. A method according to claim 25 or 26, characterized in that the reaction temperature is of from 15 to 402C.
28. A method according to claim 25 or 26, characterized in that the reaction time is of from 1 to 18 hours.
29. A method according to any one of the claims from 25 to 28, characterized in that the diluent is an aprotic diluent selected from the group comprising toluene, dimethylformamide and dimethylsulphoxide.
30. An intermediate of formula (II)
(ID
where
Rc is hydroxy, amino, di-(CrC3)alkyl-amino, tri-(CrC3)alkyl- ammoniomethyl, nitro, trifluoromethyl, nitrile, CH3C(0)NH, CH3S02NH, CH3SO2, R'R"NS02, where R' and R" are H, or linear or branched C-ι-C-6 alkyl, Rd is H, hydroxy, amino, di-(CrC3)alkyl-amino, tri-(d- C3)alkylammoniomethyl, nitro, trifluoromethyl, nitrile, CH3C(0)NH, CH3S02NH, CH3S02, R'R"NS02, where R' and R" have the meanings stated above. 31. A pharmaceutical composition containing an effective amount of a compound of formula (I):
Figure imgf000033_0001
(l)
where
X is C(0)NHCH2, NHC(O) or NHC(0)CH2; Ra is H, NH2C(0), CH3C(0)NH, CH3S02, CH3S02NH, linear or . branched C1-C3 alkyl, linear or branched C1-C3 alkoxy, or • halogen; Rb is H, linear or branched Ci-Cδ alkyl; aryl-(CrC3)alkyl optionally substituted with 1 or 2 halogen atoms, with a C1-C3 alkyl group or a C1-C3 alkoxy group; and in which a) when X is C(0)NHCH2
Rc is hydroxy, amino, di-(C-ι-C3)alkyl-amino, tri-(C C3)alkylammoniomethyl, nitro, trifluoromethyl, nitrile, CH3C(0)NH, CH3S02NH, CH3S02, R'R"NS02, where R' and R" are H, or a linear or branched CrC6 alkyl , R is H, hydroxy, amino, di-(Cι-C3)alkyl-amino, tri-(Cr C3)alkylammoniomethyl, nitro, trifluoromethyl, nitrile, CH3C(0)NH, CH3S02NH, CH3S02, R'R"NS02, where R1 and R" have the meanings stated above, with the proviso, however, that when Ra and Rd are both H, and R is isopropyl, then Rc is not hydroxy; b) when X is NHC(O) or NHC(0)CH2
Rc and Rd, which may be equal or different, are H, hydroxy, d- C-3 alkoxy, halogen, amino, di-(d-C3)alkylamino, tri-
(C-ι-C3)alkylammoniomethyl, nitro, trifluoromethyl, nitrile, CH3C(0)NH, CH3S02NH, CH3S02, R'R"NS02, where R' and R" have the meanings stated above, or of a pharmaceutically acceptable addition salt thereof with an organic or inorganic acid, and at least one pharmaceutically acceptable inert ingredient. 32. A pharmaceutical composition according to claim 31 , characterized in that it contains a compound according to any one of the preceding claims from 2 to 22.
PCT/EP2004/004390 2003-05-15 2004-04-23 Indazole having analgesic activity , Ceased WO2004101548A1 (en)

Priority Applications (17)

Application Number Priority Date Filing Date Title
JP2006529710A JP4865559B2 (en) 2003-05-15 2004-04-23 Indazole with analgesic activity
US10/549,930 US7662836B2 (en) 2003-05-15 2004-04-23 Indazole having analgesic activity
CA2519733A CA2519733C (en) 2003-05-15 2004-04-23 Indazole having analgesic activity
AT04729106T ATE501134T1 (en) 2003-05-15 2004-04-23 INDAZOLES WITH ANALGESIC ACTIVITY
AU2004238449A AU2004238449B2 (en) 2003-05-15 2004-04-23 Indazole having analgesic activity ,
UAA200509350A UA80605C2 (en) 2003-05-15 2004-04-23 The indazoles having an analgesic activity
EA200501818A EA008499B1 (en) 2003-05-15 2004-04-23 Indazole having analgesic activity
HK06107998.1A HK1087705B (en) 2003-05-15 2004-04-23 Indazoles having analgesic activity
SI200431647T SI1622892T1 (en) 2003-05-15 2004-04-23 Indazoles having analgesic activity
PL04729106T PL1622892T3 (en) 2003-05-15 2004-04-23 Indazoles having analgesic activity
KR1020057021768A KR101093048B1 (en) 2003-05-15 2004-04-23 Indazole with analgesic action
MXPA05012297A MXPA05012297A (en) 2003-05-15 2004-04-23 Indazole having analgesic activity ,.
EP04729106A EP1622892B1 (en) 2003-05-15 2004-04-23 Indazoles having analgesic activity
DK04729106.7T DK1622892T3 (en) 2003-05-15 2004-04-23 Indazoles with analgesic activity
DE602004031726T DE602004031726D1 (en) 2003-05-15 2004-04-23 INDAZOLE WITH ANALGETIC ACTIVITY
IL170769A IL170769A (en) 2003-05-15 2005-09-08 Indazoles, pharmaceutical compositions comprising them, uses thereof for the preparation of medicaments and methods of their preparation
US12/638,470 US8415476B2 (en) 2003-05-15 2009-12-15 Indazole having analgesic activity

Applications Claiming Priority (2)

Application Number Priority Date Filing Date Title
ITMI2003A000972 2003-05-15
IT000972A ITMI20030972A1 (en) 2003-05-15 2003-05-15 INDAZOLO EQUIPPED WITH ANALGESIC ACTIVITY, METHOD TO PREPARE IT AND PHARMACEUTICAL COMPOSITION THAT INCLUDES IT.

Related Child Applications (2)

Application Number Title Priority Date Filing Date
US10549930 A-371-Of-International 2004-04-23
US12/638,470 Division US8415476B2 (en) 2003-05-15 2009-12-15 Indazole having analgesic activity

Publications (1)

Publication Number Publication Date
WO2004101548A1 true WO2004101548A1 (en) 2004-11-25

Family

ID=33446432

Family Applications (1)

Application Number Title Priority Date Filing Date
PCT/EP2004/004390 Ceased WO2004101548A1 (en) 2003-05-15 2004-04-23 Indazole having analgesic activity ,

Country Status (21)

Country Link
US (2) US7662836B2 (en)
EP (1) EP1622892B1 (en)
JP (1) JP4865559B2 (en)
KR (1) KR101093048B1 (en)
CN (1) CN100364988C (en)
AR (1) AR044317A1 (en)
AT (1) ATE501134T1 (en)
AU (1) AU2004238449B2 (en)
CA (1) CA2519733C (en)
DE (1) DE602004031726D1 (en)
DK (1) DK1622892T3 (en)
EA (1) EA008499B1 (en)
ES (1) ES2359242T3 (en)
GE (1) GEP20074183B (en)
IL (1) IL170769A (en)
IT (1) ITMI20030972A1 (en)
MX (1) MXPA05012297A (en)
PL (1) PL1622892T3 (en)
SI (1) SI1622892T1 (en)
UA (1) UA80605C2 (en)
WO (1) WO2004101548A1 (en)

Cited By (7)

* Cited by examiner, † Cited by third party
Publication number Priority date Publication date Assignee Title
WO2008061688A1 (en) 2006-11-22 2008-05-29 Aziende Chimiche Riunite Angelini Francesco A.C.R.A.F. S.P.A. 2 -alkyl- indazole compounds for the treatment of certain cns-related disorders
JP2009513544A (en) * 2003-07-18 2009-04-02 アジェンデ・キミケ・リウニテ・アンジェリニ・フランチェスコ・ア・チ・エレ・ア・エフェ・ソシエタ・ペル・アチオニ Methods of using indazole derivatives for the treatment of neuropathic pain
WO2009062883A1 (en) * 2007-11-12 2009-05-22 Aziende Chimiche Riunite Angelini Francesco A.C.R.A.F. S.P.A. Drug active in neuropathic pain
WO2013124158A1 (en) 2012-02-21 2013-08-29 Aziende Chimiche Riunite Angelini Francesco A.C.R.A.F. S.P.A. 1h-indazole-3-carboxamide compounds as glycogen synthase kinase 3 beta inhibitors
WO2013124169A1 (en) 2012-02-21 2013-08-29 Aziende Chimiche Riunite Angelini Francesco A.C.R.A.F. S.P.A. Use of 1h-indazole-3-carboxamide compounds as glycogen synthase kinase 3 beta inhibitors
US8686147B2 (en) 2008-07-29 2014-04-01 Aziende Chimiche Riunite Angelini Francesco A.C.R.A.F. S.P.A. Compound with serotoninergic activity, process for preparing it and pharmaceutical composition comprising it
WO2019215075A1 (en) 2018-05-07 2019-11-14 Aziende Chimiche Riunite Angelini Francesco - A.C.R.A.F. S.P.A. 1h-indazole-3-carboxamide compounds as glycogen synthase kinase 3 beta inhibitors

Citations (3)

* Cited by examiner, † Cited by third party
Publication number Priority date Publication date Assignee Title
WO1998046589A2 (en) * 1997-04-15 1998-10-22 Aziende Chimiche Riunite Angelini Francesco A.C.R.A.F. S.P.A. Indazole amide compounds as serotoninergic agents
EP0908459A1 (en) * 1997-10-07 1999-04-14 Eli Lilly And Company 5-HT4 Agonists and antagonists
WO2003004026A1 (en) * 2001-07-05 2003-01-16 Grünenthal GmbH Substituted 1-phenethylpiperidine compounds used as inter alia analgesics

Family Cites Families (7)

* Cited by examiner, † Cited by third party
Publication number Priority date Publication date Assignee Title
US6017931A (en) 1994-03-01 2000-01-25 Fmc Corporation Insecticidal compositions containing n-(substituted phenylmethyl)-4-[bis(substituted phenyl)methyl]piperidines
US5654320A (en) * 1995-03-16 1997-08-05 Eli Lilly And Company Indazolecarboxamides
KR19990022096A (en) * 1995-05-31 1999-03-25 쇼다 오사무 Indazole derivatives with monocyclic amino groups
CN1093801C (en) * 1995-11-08 2002-11-06 东丽株式会社 Direct drawing type waterless planographic original form plate
TR199900321T2 (en) 1996-08-16 1999-04-21 Smithkline Beecham P.L.C. N-(1-nbutil - 4 - piperidil)metil) -3,4-dihidro -2H- (1,3) oksazino (3.2A) indol-10- Karboksamit haz�rlama prosesi ve prosesteki tuzlar ve ara �r�nler .
US6339087B1 (en) * 1997-08-18 2002-01-15 Syntex (U.S.A.) Llc Cyclic amine derivatives-CCR-3 receptor antagonists
AU2002353186A1 (en) * 2001-12-19 2003-06-30 Smithkline Beecham P.L.C. (1-h-indazol-3-yl) -amide derivatives as gsk-3 inhibitors

Patent Citations (3)

* Cited by examiner, † Cited by third party
Publication number Priority date Publication date Assignee Title
WO1998046589A2 (en) * 1997-04-15 1998-10-22 Aziende Chimiche Riunite Angelini Francesco A.C.R.A.F. S.P.A. Indazole amide compounds as serotoninergic agents
EP0908459A1 (en) * 1997-10-07 1999-04-14 Eli Lilly And Company 5-HT4 Agonists and antagonists
WO2003004026A1 (en) * 2001-07-05 2003-01-16 Grünenthal GmbH Substituted 1-phenethylpiperidine compounds used as inter alia analgesics

Cited By (25)

* Cited by examiner, † Cited by third party
Publication number Priority date Publication date Assignee Title
JP4881729B2 (en) * 2003-07-18 2012-02-22 アジェンデ・キミケ・リウニテ・アンジェリニ・フランチェスコ・ア・チ・エレ・ア・エフェ・ソシエタ・ペル・アチオニ Methods of using indazole derivatives for the treatment of neuropathic pain
JP2009513544A (en) * 2003-07-18 2009-04-02 アジェンデ・キミケ・リウニテ・アンジェリニ・フランチェスコ・ア・チ・エレ・ア・エフェ・ソシエタ・ペル・アチオニ Methods of using indazole derivatives for the treatment of neuropathic pain
US20100056573A1 (en) * 2006-11-22 2010-03-04 Aziende Chimiche Riunite Angelini Francesco A.C.R.A.F. S.P.A. 2-alkyl-indazole compounds for the treatment of certain cns-related disorders
JP2010510265A (en) * 2006-11-22 2010-04-02 アジェンデ・キミケ・リウニテ・アンジェリニ・フランチェスコ・ア・チ・エレ・ア・エフェ・ソシエタ・ペル・アチオニ 2-alkyl-indazole compounds for the treatment of certain CNS related disorders
US8507528B2 (en) 2006-11-22 2013-08-13 Aziende Chimiche Riunite Angelini Francesco A.C.R.A.F. S.P.A. 2-alkyl-indazole compounds for the treatment of certain cns-related disorders
WO2008061688A1 (en) 2006-11-22 2008-05-29 Aziende Chimiche Riunite Angelini Francesco A.C.R.A.F. S.P.A. 2 -alkyl- indazole compounds for the treatment of certain cns-related disorders
AU2008323026B2 (en) * 2007-11-12 2014-06-19 Aziende Chimiche Riunite Angelini Francesco A.C.R.A.F. S.P.A. Drug active in neuropathic pain
JP2011503030A (en) * 2007-11-12 2011-01-27 アジェンデ・キミケ・リウニテ・アンジェリニ・フランチェスコ・ア・チ・エレ・ア・エフェ・ソシエタ・ペル・アチオニ Drugs active against neuropathic pain
WO2009062883A1 (en) * 2007-11-12 2009-05-22 Aziende Chimiche Riunite Angelini Francesco A.C.R.A.F. S.P.A. Drug active in neuropathic pain
US8455519B2 (en) 2007-11-12 2013-06-04 Aziende Chimiche Riunite Angelini Francesco A.C.R.A.F. S.P.A. Drug active in neuropathic pain
KR101555626B1 (en) 2007-11-12 2015-09-24 아지엔드 키미쉐 리유나이트 안젤리니 프란체스코 에이.씨.알.에이.에프. 에스.피.에이 Drug active in neuropathic pain
CN101855220B (en) * 2007-11-12 2014-07-30 方济各安吉利克化学联合股份有限公司 Drugs active in neuropathic pain
EA019405B1 (en) * 2007-11-12 2014-03-31 Ацьенде Кимике Рьюните Анджелини Франческо A.К.P.A.Ф. С.П.А. Drug active in neuropathic pain
US8686147B2 (en) 2008-07-29 2014-04-01 Aziende Chimiche Riunite Angelini Francesco A.C.R.A.F. S.P.A. Compound with serotoninergic activity, process for preparing it and pharmaceutical composition comprising it
WO2013124169A1 (en) 2012-02-21 2013-08-29 Aziende Chimiche Riunite Angelini Francesco A.C.R.A.F. S.P.A. Use of 1h-indazole-3-carboxamide compounds as glycogen synthase kinase 3 beta inhibitors
CN104080781A (en) * 2012-02-21 2014-10-01 方济各安吉利克化学联合股份有限公司 Use of 1H-indazole-3-carboxamide compounds as glycogen synthase kinase 3 beta inhibitors
KR20140122725A (en) * 2012-02-21 2014-10-20 아지엔드 키미쉐 리유나이트 안젤리니 프란체스코 에이.씨.알.에이.에프. 에스.피.에이 Use of 1H-Indazole-3-Carboxamide Compounds as Glycogen Synthase Kinase 3 Beta Inhibitors
WO2013124158A1 (en) 2012-02-21 2013-08-29 Aziende Chimiche Riunite Angelini Francesco A.C.R.A.F. S.P.A. 1h-indazole-3-carboxamide compounds as glycogen synthase kinase 3 beta inhibitors
US9163013B2 (en) 2012-02-21 2015-10-20 Aziende Chimiche Riunite Angelini Francesco A.C.R.A.F. S.P.A. Use of 1H-indazole-3-carboxamide compounds as glycogen synthase kinase 3 beta inhibitors
CN104080781B (en) * 2012-02-21 2016-05-11 方济各安吉利克化学联合股份有限公司 1H-indazole-3-benzamide compound is as the purposes of glycogen synthase kinase 3-beta inhibitors
AU2013224313B2 (en) * 2012-02-21 2017-03-30 Aziende Chimiche Riunite Angelini Francesco A.C.R.A.F. S.P.A. Use of 1H-indazole-3-carboxamide compounds as glycogen synthase kinase 3 beta inhibitors
EA026952B1 (en) * 2012-02-21 2017-06-30 Ацьенде Кимике Рьюните Анджелини Франческо А.К.Р.А.Ф. С.П.А. Use of 1h-indazole-3-carboxamide compounds as glycogen synthase kinase 3 beta inhibitors
US9700551B2 (en) 2012-02-21 2017-07-11 Aziende Chimiche Riunite Angelini Francesco A.C.R.A.F. S.P.A. Use of 1H-indazole-3-carboxamide compounds as glycogen synthase kinase 3 beta inhibitors
KR102101702B1 (en) 2012-02-21 2020-04-20 아지엔드 키미쉐 리유나이트 안젤리니 프란체스코 에이.씨.알.에이.에프. 에스.피.에이 Use of 1H-Indazole-3-Carboxamide Compounds as Glycogen Synthase Kinase 3 Beta Inhibitors
WO2019215075A1 (en) 2018-05-07 2019-11-14 Aziende Chimiche Riunite Angelini Francesco - A.C.R.A.F. S.P.A. 1h-indazole-3-carboxamide compounds as glycogen synthase kinase 3 beta inhibitors

Also Published As

Publication number Publication date
JP4865559B2 (en) 2012-02-01
ATE501134T1 (en) 2011-03-15
EP1622892A1 (en) 2006-02-08
PL1622892T3 (en) 2011-08-31
IL170769A (en) 2012-05-31
GEP20074183B (en) 2007-08-10
DK1622892T3 (en) 2011-06-27
CA2519733A1 (en) 2004-11-25
KR20060009937A (en) 2006-02-01
US7662836B2 (en) 2010-02-16
JP2006528212A (en) 2006-12-14
SI1622892T1 (en) 2011-06-30
ITMI20030972A1 (en) 2004-11-16
KR101093048B1 (en) 2011-12-13
HK1087705A1 (en) 2006-10-20
AU2004238449B2 (en) 2010-09-02
AU2004238449A1 (en) 2004-11-25
US8415476B2 (en) 2013-04-09
DE602004031726D1 (en) 2011-04-21
IL170769A0 (en) 2009-02-11
MXPA05012297A (en) 2006-01-30
US20070010555A1 (en) 2007-01-11
CN100364988C (en) 2008-01-30
UA80605C2 (en) 2007-10-10
US20100094015A1 (en) 2010-04-15
EP1622892B1 (en) 2011-03-09
EA200501818A1 (en) 2006-06-30
CN1777597A (en) 2006-05-24
CA2519733C (en) 2011-11-29
EA008499B1 (en) 2007-06-29
ES2359242T3 (en) 2011-05-19
AR044317A1 (en) 2005-09-07

Similar Documents

Publication Publication Date Title
US8519139B2 (en) Indazolamides with analgesic activity
US8415476B2 (en) Indazole having analgesic activity
BG63380B1 (en) 4-substituted piperidine analogues and their application as selective antagonists of the nmda receptor subspecies
AU2007287338A1 (en) Piperidine derivatives
EP2276738A1 (en) Ether benzylidene piperidine 5-membered aryl carboxamide compounds useful as faah inhibitors
HUE031676T2 (en) Drug active in neuropathic pain
WO2019225625A1 (en) Readthrough inducer and pharmaceutical use thereof
HK1087705B (en) Indazoles having analgesic activity
WO2005013989A1 (en) Use of indazole derivatives for the treatment of neuropathic pain
HK1079199B (en) Indazolamides with analgesic activity

Legal Events

Date Code Title Description
AK Designated states

Kind code of ref document: A1

Designated state(s): AE AG AL AM AT AU AZ BA BB BG BR BW BY BZ CA CH CN CO CR CU CZ DE DK DM DZ EC EE EG ES FI GB GD GE GH GM HR HU ID IL IN IS JP KE KG KP KR KZ LC LK LR LS LT LU LV MA MD MG MK MN MW MX MZ NA NI NO NZ OM PG PH PL PT RO RU SC SD SE SG SK SL SY TJ TM TN TR TT TZ UA UG US UZ VC VN YU ZA ZM ZW

AL Designated countries for regional patents

Kind code of ref document: A1

Designated state(s): BW GH GM KE LS MW MZ SD SL SZ TZ UG ZM ZW AM AZ BY KG KZ MD RU TJ TM AT BE BG CH CY CZ DE DK EE ES FI FR GB GR HU IE IT LU MC NL PL PT RO SE SI SK TR BF BJ CF CG CI CM GA GN GQ GW ML MR NE SN TD TG

121 Ep: the epo has been informed by wipo that ep was designated in this application
DPEN Request for preliminary examination filed prior to expiration of 19th month from priority date (pct application filed from 20040101)
WWE Wipo information: entry into national phase

Ref document number: 170769

Country of ref document: IL

WWE Wipo information: entry into national phase

Ref document number: 2004238449

Country of ref document: AU

WWE Wipo information: entry into national phase

Ref document number: 2004729106

Country of ref document: EP

WWE Wipo information: entry into national phase

Ref document number: 2007010555

Country of ref document: US

Ref document number: 2519733

Country of ref document: CA

Ref document number: 10549930

Country of ref document: US

ENP Entry into the national phase

Ref document number: 2004238449

Country of ref document: AU

Date of ref document: 20040423

Kind code of ref document: A

WWP Wipo information: published in national office

Ref document number: 2004238449

Country of ref document: AU

WWE Wipo information: entry into national phase

Ref document number: 20048104363

Country of ref document: CN

WWE Wipo information: entry into national phase

Ref document number: PA/a/2005/012297

Country of ref document: MX

Ref document number: 2006529710

Country of ref document: JP

Ref document number: 1020057021768

Country of ref document: KR

WWE Wipo information: entry into national phase

Ref document number: 9111

Country of ref document: GE

Ref document number: 20050425

Country of ref document: UZ

Ref document number: 200501818

Country of ref document: EA

WWP Wipo information: published in national office

Ref document number: 1020057021768

Country of ref document: KR

WWP Wipo information: published in national office

Ref document number: 2004729106

Country of ref document: EP

WWP Wipo information: published in national office

Ref document number: 10549930

Country of ref document: US