WO2007147822A1 - Phenylpentadienoyl derivatives and their use as par 1 antagonists - Google Patents
Phenylpentadienoyl derivatives and their use as par 1 antagonists Download PDFInfo
- Publication number
- WO2007147822A1 WO2007147822A1 PCT/EP2007/056078 EP2007056078W WO2007147822A1 WO 2007147822 A1 WO2007147822 A1 WO 2007147822A1 EP 2007056078 W EP2007056078 W EP 2007056078W WO 2007147822 A1 WO2007147822 A1 WO 2007147822A1
- Authority
- WO
- WIPO (PCT)
- Prior art keywords
- phenyl
- penta
- piperazin
- dien
- general formula
- Prior art date
- Legal status (The legal status is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the status listed.)
- Ceased
Links
- 0 C*(CC(C)(C)N(CC(*)CCN)CC1)CN1C(C=CC=Cc1c(*)cccc1*)=O Chemical compound C*(CC(C)(C)N(CC(*)CCN)CC1)CN1C(C=CC=Cc1c(*)cccc1*)=O 0.000 description 1
- RWAGJRSRCSOWHV-XFRMRLMGSA-N O=C(/C=C/C=C/c(c(F)ccc1)c1F)N(CC1)CCN1c1ccccc1 Chemical compound O=C(/C=C/C=C/c(c(F)ccc1)c1F)N(CC1)CCN1c1ccccc1 RWAGJRSRCSOWHV-XFRMRLMGSA-N 0.000 description 1
Classifications
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D295/00—Heterocyclic compounds containing polymethylene-imine rings with at least five ring members, 3-azabicyclo [3.2.2] nonane, piperazine, morpholine or thiomorpholine rings, having only hydrogen atoms directly attached to the ring carbon atoms
- C07D295/16—Heterocyclic compounds containing polymethylene-imine rings with at least five ring members, 3-azabicyclo [3.2.2] nonane, piperazine, morpholine or thiomorpholine rings, having only hydrogen atoms directly attached to the ring carbon atoms acylated on ring nitrogen atoms
- C07D295/18—Heterocyclic compounds containing polymethylene-imine rings with at least five ring members, 3-azabicyclo [3.2.2] nonane, piperazine, morpholine or thiomorpholine rings, having only hydrogen atoms directly attached to the ring carbon atoms acylated on ring nitrogen atoms by radicals derived from carboxylic acids, or sulfur or nitrogen analogues thereof
- C07D295/182—Radicals derived from carboxylic acids
- C07D295/192—Radicals derived from carboxylic acids from aromatic carboxylic acids
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D241/00—Heterocyclic compounds containing 1,4-diazine or hydrogenated 1,4-diazine rings
- C07D241/02—Heterocyclic compounds containing 1,4-diazine or hydrogenated 1,4-diazine rings not condensed with other rings
- C07D241/10—Heterocyclic compounds containing 1,4-diazine or hydrogenated 1,4-diazine rings not condensed with other rings having three double bonds between ring members or between ring members and non-ring members
- C07D241/14—Heterocyclic compounds containing 1,4-diazine or hydrogenated 1,4-diazine rings not condensed with other rings having three double bonds between ring members or between ring members and non-ring members with hetero atoms or with carbon atoms having three bonds to hetero atoms with at the most one bond to halogen, e.g. ester or nitrile radicals, directly attached to ring carbon atoms
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61K—PREPARATIONS FOR MEDICAL, DENTAL OR TOILETRY PURPOSES
- A61K31/00—Medicinal preparations containing organic active ingredients
- A61K31/33—Heterocyclic compounds
- A61K31/395—Heterocyclic compounds having nitrogen as a ring hetero atom, e.g. guanethidine or rifamycins
- A61K31/495—Heterocyclic compounds having nitrogen as a ring hetero atom, e.g. guanethidine or rifamycins having six-membered rings with two or more nitrogen atoms as the only ring heteroatoms, e.g. piperazine or tetrazines
- A61K31/496—Non-condensed piperazines containing further heterocyclic rings, e.g. rifampin, thiothixene or sparfloxacin
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P1/00—Drugs for disorders of the alimentary tract or the digestive system
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P1/00—Drugs for disorders of the alimentary tract or the digestive system
- A61P1/16—Drugs for disorders of the alimentary tract or the digestive system for liver or gallbladder disorders, e.g. hepatoprotective agents, cholagogues, litholytics
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P11/00—Drugs for disorders of the respiratory system
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P29/00—Non-central analgesic, antipyretic or antiinflammatory agents, e.g. antirheumatic agents; Non-steroidal antiinflammatory drugs [NSAID]
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P43/00—Drugs for specific purposes, not provided for in groups A61P1/00-A61P41/00
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P7/00—Drugs for disorders of the blood or the extracellular fluid
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P7/00—Drugs for disorders of the blood or the extracellular fluid
- A61P7/02—Antithrombotic agents; Anticoagulants; Platelet aggregation inhibitors
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P9/00—Drugs for disorders of the cardiovascular system
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P9/00—Drugs for disorders of the cardiovascular system
- A61P9/04—Inotropic agents, i.e. stimulants of cardiac contraction; Drugs for heart failure
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P9/00—Drugs for disorders of the cardiovascular system
- A61P9/10—Drugs for disorders of the cardiovascular system for treating ischaemic or atherosclerotic diseases, e.g. antianginal drugs, coronary vasodilators, drugs for myocardial infarction, retinopathy, cerebrovascula insufficiency, renal arteriosclerosis
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P9/00—Drugs for disorders of the cardiovascular system
- A61P9/12—Antihypertensives
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D213/00—Heterocyclic compounds containing six-membered rings, not condensed with other rings, with one nitrogen atom as the only ring hetero atom and three or more double bonds between ring members or between ring members and non-ring members
- C07D213/02—Heterocyclic compounds containing six-membered rings, not condensed with other rings, with one nitrogen atom as the only ring hetero atom and three or more double bonds between ring members or between ring members and non-ring members having three double bonds between ring members or between ring members and non-ring members
- C07D213/04—Heterocyclic compounds containing six-membered rings, not condensed with other rings, with one nitrogen atom as the only ring hetero atom and three or more double bonds between ring members or between ring members and non-ring members having three double bonds between ring members or between ring members and non-ring members having no bond between the ring nitrogen atom and a non-ring member or having only hydrogen or carbon atoms directly attached to the ring nitrogen atom
- C07D213/60—Heterocyclic compounds containing six-membered rings, not condensed with other rings, with one nitrogen atom as the only ring hetero atom and three or more double bonds between ring members or between ring members and non-ring members having three double bonds between ring members or between ring members and non-ring members having no bond between the ring nitrogen atom and a non-ring member or having only hydrogen or carbon atoms directly attached to the ring nitrogen atom with hetero atoms or with carbon atoms having three bonds to hetero atoms with at the most one bond to halogen, e.g. ester or nitrile radicals, directly attached to ring carbon atoms
- C07D213/61—Halogen atoms or nitro radicals
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D213/00—Heterocyclic compounds containing six-membered rings, not condensed with other rings, with one nitrogen atom as the only ring hetero atom and three or more double bonds between ring members or between ring members and non-ring members
- C07D213/02—Heterocyclic compounds containing six-membered rings, not condensed with other rings, with one nitrogen atom as the only ring hetero atom and three or more double bonds between ring members or between ring members and non-ring members having three double bonds between ring members or between ring members and non-ring members
- C07D213/04—Heterocyclic compounds containing six-membered rings, not condensed with other rings, with one nitrogen atom as the only ring hetero atom and three or more double bonds between ring members or between ring members and non-ring members having three double bonds between ring members or between ring members and non-ring members having no bond between the ring nitrogen atom and a non-ring member or having only hydrogen or carbon atoms directly attached to the ring nitrogen atom
- C07D213/60—Heterocyclic compounds containing six-membered rings, not condensed with other rings, with one nitrogen atom as the only ring hetero atom and three or more double bonds between ring members or between ring members and non-ring members having three double bonds between ring members or between ring members and non-ring members having no bond between the ring nitrogen atom and a non-ring member or having only hydrogen or carbon atoms directly attached to the ring nitrogen atom with hetero atoms or with carbon atoms having three bonds to hetero atoms with at the most one bond to halogen, e.g. ester or nitrile radicals, directly attached to ring carbon atoms
- C07D213/72—Nitrogen atoms
- C07D213/74—Amino or imino radicals substituted by hydrocarbon or substituted hydrocarbon radicals
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D241/00—Heterocyclic compounds containing 1,4-diazine or hydrogenated 1,4-diazine rings
- C07D241/02—Heterocyclic compounds containing 1,4-diazine or hydrogenated 1,4-diazine rings not condensed with other rings
- C07D241/04—Heterocyclic compounds containing 1,4-diazine or hydrogenated 1,4-diazine rings not condensed with other rings having no double bonds between ring members or between ring members and non-ring members
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D295/00—Heterocyclic compounds containing polymethylene-imine rings with at least five ring members, 3-azabicyclo [3.2.2] nonane, piperazine, morpholine or thiomorpholine rings, having only hydrogen atoms directly attached to the ring carbon atoms
- C07D295/04—Heterocyclic compounds containing polymethylene-imine rings with at least five ring members, 3-azabicyclo [3.2.2] nonane, piperazine, morpholine or thiomorpholine rings, having only hydrogen atoms directly attached to the ring carbon atoms with substituted hydrocarbon radicals attached to ring nitrogen atoms
- C07D295/10—Heterocyclic compounds containing polymethylene-imine rings with at least five ring members, 3-azabicyclo [3.2.2] nonane, piperazine, morpholine or thiomorpholine rings, having only hydrogen atoms directly attached to the ring carbon atoms with substituted hydrocarbon radicals attached to ring nitrogen atoms substituted by doubly bound oxygen or sulphur atoms
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D295/00—Heterocyclic compounds containing polymethylene-imine rings with at least five ring members, 3-azabicyclo [3.2.2] nonane, piperazine, morpholine or thiomorpholine rings, having only hydrogen atoms directly attached to the ring carbon atoms
- C07D295/04—Heterocyclic compounds containing polymethylene-imine rings with at least five ring members, 3-azabicyclo [3.2.2] nonane, piperazine, morpholine or thiomorpholine rings, having only hydrogen atoms directly attached to the ring carbon atoms with substituted hydrocarbon radicals attached to ring nitrogen atoms
- C07D295/10—Heterocyclic compounds containing polymethylene-imine rings with at least five ring members, 3-azabicyclo [3.2.2] nonane, piperazine, morpholine or thiomorpholine rings, having only hydrogen atoms directly attached to the ring carbon atoms with substituted hydrocarbon radicals attached to ring nitrogen atoms substituted by doubly bound oxygen or sulphur atoms
- C07D295/112—Heterocyclic compounds containing polymethylene-imine rings with at least five ring members, 3-azabicyclo [3.2.2] nonane, piperazine, morpholine or thiomorpholine rings, having only hydrogen atoms directly attached to the ring carbon atoms with substituted hydrocarbon radicals attached to ring nitrogen atoms substituted by doubly bound oxygen or sulphur atoms with the ring nitrogen atoms and the doubly bound oxygen or sulfur atoms separated by carbocyclic rings or by carbon chains interrupted by carbocyclic rings
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D333/00—Heterocyclic compounds containing five-membered rings having one sulfur atom as the only ring hetero atom
- C07D333/02—Heterocyclic compounds containing five-membered rings having one sulfur atom as the only ring hetero atom not condensed with other rings
- C07D333/04—Heterocyclic compounds containing five-membered rings having one sulfur atom as the only ring hetero atom not condensed with other rings not substituted on the ring sulphur atom
- C07D333/26—Heterocyclic compounds containing five-membered rings having one sulfur atom as the only ring hetero atom not condensed with other rings not substituted on the ring sulphur atom with hetero atoms or with carbon atoms having three bonds to hetero atoms with at the most one bond to halogen, e.g. ester or nitrile radicals, directly attached to ring carbon atoms
- C07D333/30—Hetero atoms other than halogen
- C07D333/36—Nitrogen atoms
Definitions
- the present invention relates to phenylpentadienoyl derivatives, a method of manufacturing same, pharmaceutical compositions comprised of same and the use of same as drugs for the treatment and/or prevention of arterial and venous thrombosis, acute coronary syndromes, restenosis, stable angina, heart rhythm disorders, myocardial infarction, hypertension, heart failure, stroke, inflammatory disorders, pulmonary diseases, gastrointestinal diseases, fibrosis development in chronic liver disease patients, cancer and skin diseases.
- the present invention also relates to combinations of the inventive compounds with other cardiovascular agents.
- Thrombosis is regarded as a primary factor in vascular occlusion, which is the cause of a number of pathophysiological complications. Antithrombotic therapy is thus extremely important as it can reduce the risk of cardiovascular mortality and coronary events. Although several types of molecules have shown effective antithrombotic activity in man, there remains a need for novel molecules that provide advantages compared to existing compounds, some of which have a negative impact on bleeding time or are accompanied by other undesirable side effects (such as, for example, the risk of ulcer with aspirin) .
- Protease-activated receptor-1 was recently cloned (Vu et al., Cell, 1991, 64: 1057-1068) and its mechanism of action elucidated (Coughlin et al., J. Clin. Invest. 1992, 89(2): 351-355).
- This receptor notably present on the surface of platelets but also on the surface of endothelial cells (O'Brien et al . , J. Biol. Chem. 2000, 275: 13502-13509), smooth muscle cells (Hamilton et al., Br. J. Pharmacol. 2000, 130: 181-188) and fibroblasts (Hung et al., J. Cell. Biol.
- thrombin receptor is activated by thrombin and thus is also called thrombin receptor.
- the N-terminus of the protein is cleaved by thrombin between arginine 41 and serine 42 to free a new end which will act, after folding upon the active site, as a receptor agonist (Vu et al., Nature, 1991, 353, 674-677).
- this specific PAR-I receptor activation mechanism leads to thrombin-mediated platelet aggregation.
- the blocking of this activation can inhibit thrombin-mediated platelet aggregation (Ahn et al . , Drug of the Future, 2001, 26: 1065- 1085) .
- the blocking of these receptors can thus lead to the treatment or prevention of thrombosis (Derian et al., J. Pharmacol. Exp. Ther., 2003, 855-861), acute coronary syndromes (Ossovskaya et al., Physiol. Rev., 2004, 84: 579-
- Cardiovasc. Hematol. Agents., 2003, 13-36 can reduce myocardial necroses during infarction or reperfusion
- PAR-I antagonist activity can prevent certain inflammatory diseases (Moffatt et al., Curr. Op. Pharmacol., 2004, 221-229) .
- PAR-I receptor antagonist activity can prevent certain inflammatory diseases (Vergnolle et al., J. Clin. Invest., 2004, 1444-1456).
- PAR-I antagonists can also be of use in the treatment of fibroses in patients with chronic liver disease (Fiorucci et al., Hepatology, 2004, 39: 365-375) .
- PAR-I antagonists can be of interest in dermatology to treat certain skin diseases (Schechter et al., J. Cell. Physiol., 1998, 176:365-373; Algermissen et al., Arch. Dermatol. Res., 2000, 292:488-495; Meyer-Hoffert et al., Exp. Dermatol., 2004, 13: 234-241).
- the present invention relates to a novel class of PAR-I antagonists that are distinguished from the prior art by their different chemical structure and their remarkable biological property.
- Compounds of the present invention are of general formula (D :
- Ri and R 2 identical or different, represent: an atom of hydrogen or halogen, CN or NO2, with R 1 and R2 not representing hydrogen simultaneously, m represents :
- R3 represents: phenyl substituted or not by one or more residues chosen among halogen, hydroxyl or Ci-C ⁇ alkyl; C2-C6 alkyl substituted or not by one or more residues chosen among halogen or hydroxyl; cycloalkyl; pyridine; thiophene; pyrrole substituted or not by Ci-C ⁇ alkyl; thiazole or furan; or the therapeutically-acceptable salts or solvates thereof.
- halogen represents fluorine, chlorine, bromine or iodine.
- alkyl represents linear or branched, saturated or unsaturated aliphatic hydrocarbon chains comprising the specified number of carbon atoms.
- cycloalkyl represents cyclic hydrocarbon chains comprising 3 to 10 carbon atoms.
- Therapeutically-acceptable salts of compounds of the present invention include conventional nontoxic salts of compounds of the invention such as those formed from organic or inorganic acids.
- inorganic acid salts such as hydrochloric, hydrobromic, phosphoric and sulfuric acids
- organic acid salts such as acetic, trifluoroacetic, propionic, succinic, fumaric, malic, tartaric, citric, ascorbic, maleic, glutamic, benzoic, salicylic, toluenesulfonic, methanesulfonic, stearic and lactic acids.
- salts can be synthesized from compounds of the invention containing a base moiety and corresponding acids according to conventional chemical methods.
- Therapeutically-acceptable solvates of compounds of the present invention include conventional solvates such as those formed during the final preparation step of compounds of the invention as a result of the presence of solvents. Solvates due to the presence of water or ethanol can be cited as an example .
- one particularly advantageous class of compounds are compounds of general formula (I) wherein R 1 is nitro, R-2 is hydrogen, m equals 1, n equals 0 and R3 is phenyl substituted by one or more halogens or Ci-C ⁇ alkyls, cycloalkyl or pyridine.
- a second particularly advantageous class of compounds corresponds to compounds of general formula (I) wherein Ri is cyano, R2 is hydrogen, m equals 1, n equals 0 and R3 is phenyl substituted by one or more halogens or Ci-C ⁇ alkyls, cycloalkyl or pyridine.
- the present invention also relates to the preparation of compounds of general formula (I) by the general methods described in the following synthesis diagrams supplemented by, as the case may be, any standard technique described in the literature, known to those persons skilled in the art, or presented in the experiments section.
- Diagram 1 illustrates the first general method that can be used for the preparation of compounds of general formula
- Ri, R2 and R3 are defined as in the preceding description of general formula (I) .
- n represents 1 or 2 only
- Pi represents a protective group
- X can represent a group such as chlorine or hydroxyl .
- the starting compound, of general formula (II) can be prepared by methods and techniques known to those persons skilled in the art.
- a particularly advantageous method consists of reacting benzyl halide with triphenylphosphine in a polar solvent such as DMF or DMSO at a temperature between 20 0 C and 100 0 C to form phosphonium salt.
- Phosphonium salt can then be deprotonated using a base such as NaH, for example, in a solvent such as DMF or THF at a temperature between -20 0 C and 40 0 C and then reacting with an ⁇ , ⁇ -unsaturated aldehyde carrying an ester such as, for example, ethyl (2E) -4-oxobut-2-enoate .
- a base such as NaH
- a solvent such as DMF or THF at a temperature between -20 0 C and 40 0 C and then reacting with an ⁇ , ⁇ -unsaturated aldehyde carrying an ester such as, for example, ethyl (2E) -4-oxobut-2-enoate .
- the ester obtained (mixture of Z/E and E/E isomers) is first isomerized by treatment with iodine in a polar solvent such as acetonitrile to yield the E/E isomer exclusively, and then saponified by treatment with a mineral base such as KOH, NaOH or LiOH in a polar solvent such as water, ethanol or THF at a temperature between 20 0 C and 100 0 C, yielding compounds (II) in which X would thus be hydroxyl.
- a mineral base such as KOH, NaOH or LiOH
- a polar solvent such as water, ethanol or THF
- a second particularly advantageous method consists of reacting an aromatic aldehyde with a phosphonate such as ethyl 4- (diethoxyphosphoryl) but-2-enoate in the presence of a base such as, for example, NaH, CS2CO3 or K2CO3 in a solvent such as THF, dichloromethane or dichloroethane at a temperature between -20 0 C and 100 0 C.
- a phosphonate such as ethyl 4- (diethoxyphosphoryl) but-2-enoate
- a base such as, for example, NaH, CS2CO3 or K2CO3
- a solvent such as THF, dichloromethane or dichloroethane
- the ester obtained (mixture of Z/E and E/E isomers) is first isomerized by treatment with iodine in a polar solvent such as acetonitrile, yielding the E/E isomer exclusively, and then saponified by treatment with a mineral base such as KOH, NaOH or LiOH in a polar solvent such as water, ethanol or THF at a temperature between 20 0 C and 100 0 C, yielding compounds (II) in which X would thus be hydroxyl .
- a mineral base such as KOH, NaOH or LiOH
- a polar solvent such as water, ethanol or THF
- a third particularly advantageous method consists of reacting an aromatic ⁇ , ⁇ - unsaturated aldehyde with ethyl (diethoxyphosphoryl) acetate in the presence of a base such as, for example, NaH, CS2CO3 or K2CO3 in a solvent such as THF, dichloromethane or dichloroethane at a temperature between -20 0 C and 100 0 C.
- a base such as, for example, NaH, CS2CO3 or K2CO3
- a solvent such as THF, dichloromethane or dichloroethane
- the ester obtained can be saponified by treatment with a mineral base such as KOH, NaOH or LiOH in a solvent such as water, ethanol or THF at a temperature between 20 0 C and 100 0 C, yielding compounds (II) in which X would thus be hydroxyl.
- a fourth particularly advantageous method consists of reacting an aromatic carrying a halogen, such as bromine or iodine, with an (E) -penta-2, 4-dienoyl ester, such as methyl or ethyl (E) -penta-2, 4-dienoate ester, in the presence of a palladium catalyst such as palladium acetate, a phosphine such as tri-o- tolylphosphine or tri-phenylphosphine in the presence of a base such as, for example, Et3N or iPr 2 NEt, in an open or sealed reactor, without a solvent or with a solvent such as DMF, DMSO or DMA at a temperature between 20 0 C and 120 0 C.
- a palladium catalyst such as palladium acetate
- a phosphine such as tri-o- tolylphosphine or tri-phenylphosphine
- a base such as, for example
- the ester thus obtained (primarily the E/E isomer) can be saponified by treatment with a mineral base such as KOH, NaOH or LiOH in a solvent such as water, ethanol or THF at a temperature between 20 0 C and 100 0 C, yielding compounds (II) in which X would thus be hydroxyl.
- the first step is a condensation reaction between the carboxylic acid (II) and the amine (III) . This reaction can be carried out by methods and techniques known to those persons skilled in the art.
- a particularly advantageous method consists of reacting these two entities in the presence of l-(3- dimethylaminopropyl) -3-ethyl-carbodiimide (EDC), 3-hydroxy- 1, 2, 3-benzotriazin-4 (3H) -one, and a tertiary amine such as diisopropylethylamine in a polar aprotic solvent, such as dichloromethane, at a temperature between -15 0 C and 40 0 C.
- EDC l-(3- dimethylaminopropyl) -3-ethyl-carbodiimide
- 3-hydroxy- 1, 2, 3-benzotriazin-4 (3H) -one 3-hydroxy- 1, 2, 3-benzotriazin-4 (3H) -one
- a tertiary amine such as diisopropylethylamine
- a polar aprotic solvent such as dichloromethane
- the first step consists of the reaction between an acid chloride and an amine.
- This reaction can be carried out by methods and techniques known to those persons skilled in the art.
- a particularly advantageous method consists of reacting the two entities in the presence of an organic or inorganic base such as, for example, Et 3 N, iPr 2 NEt, pyridine, NaH, CS2CO3 or K2CO3 in a solvent such as THF, dichloromethane, DMF or DMSO at a temperature between -20° and 100 0 C.
- an organic or inorganic base such as, for example, Et 3 N, iPr 2 NEt, pyridine, NaH, CS2CO3 or K2CO3
- a solvent such as THF, dichloromethane, DMF or DMSO
- the intermediate obtained can react with a reagent of formula R 3 (CH 2 ) n Y, wherein Y represents a leaving group such as, for example, Cl, Br, I, OSO 2 CH 3 , OSO 2 CF 3 or O-tosyl.
- the reaction will be carried out in the presence of an organic or inorganic base such as, for example, Et 3 N, iPr 2 NEt, NaH, Cs 2 CO 3 or K 2 CO 3 capable of being supported on a resin such as PS-DIEA or MP-carbonate, in a polar anhydrous solvent such as dichloromethane, THF, DMF or DMSO at a temperature between -20° and 100 0 C.
- an organic or inorganic base such as, for example, Et 3 N, iPr 2 NEt, NaH, Cs 2 CO 3 or K 2 CO 3 capable of being supported on a resin such as PS-DIEA or MP-carbonate
- a polar anhydrous solvent such as dichloromethane, THF, DMF or DMSO
- Another preparation method consists of carrying out a reducing amination reaction using an aldehyde of formula R 3 - (CH 2 ) n _i-CHO in which R 3 and n are as defined previously, with the deprotected amine of general formula (IV) and a reducing agent such as NaBH 4 , NaBH 3 CN or NaBH(OAc) 3 capable of being supported on a resin such as MP-BH 3 CN, in a polar solvent such as 1,2- dichloroethane, dichloromethane, THF, DMF or MeOH, at a pH that can be controlled by the addition of an acid such as acetic acid, at a temperature between -20 0 C and 100 0 C.
- a reducing agent such as NaBH 4 , NaBH 3 CN or NaBH(OAc) 3 capable of being supported on a resin such as MP-BH 3 CN
- a polar solvent such as 1,2- dichloroethane, dichloromethane, THF, DMF or MeOH
- Diagram 2 illustrates the second general method that can be used for the preparation of compounds of general formula (I) .
- Ri, R 2 , R3 and n are defined as in the description of general formula (I) .
- X can represent a group such as chlorine or hydroxyl .
- the starting compound, of general formula (II) can be prepared by methods and techniques known to those persons skilled in the art, in particular those described above.
- synthesis consists of the reaction between an acid chloride and an amine. This reaction can be carried out by methods and techniques known to those persons skilled in the art.
- a particularly advantageous method consists of reacting the two entities in the presence of an organic or inorganic base such as, for example, Et 3 N, iPr 2 NEt, pyridine, NaH, CS 2 CO3 or K 2 CO 3 in a solvent such as THF, dichloromethane, DMF or DMSO at a temperature between -20° and 100 0 C.
- an organic or inorganic base such as, for example, Et 3 N, iPr 2 NEt, pyridine, NaH, CS 2 CO3 or K 2 CO 3 in a solvent such as THF, dichloromethane, DMF or DMSO
- a solvent such as THF, dichloromethane, DMF or DMSO
- synthesis consists of condensation between the carboxylic acid (II) and the amine (V) .
- the reaction can be carried out by methods and techniques known to those persons skilled in the art.
- a particularly advantageous method consists of condensing a carboxylic acid of general formula (II) with an amine of general formula (III) in the presence of 1- (3-dimethylaminopropyl) -3-ethyl- carbodiimide (EDC), 3-hydroxy-l, 2, 3-benzotriazin-4 (3H) -one and a tertiary amine such as diisopropylethylamine, in a polar aprotic solvent such as dichloromethane, at a temperature between -15 0 C and 40 0 C.
- EDC 1- (3-dimethylaminopropyl) -3-ethyl- carbodiimide
- 3-hydroxy-l 2, 3-benzotriazin-4 (3H) -one
- a tertiary amine such as diisopropylethylamine
- Example IA Ethyl 5- (2-cyano-phenyl) -penta-2, 4-dienoate
- the syrup obtained is purified by silica column chromatography and eluted with a 9/1 EDP/AcOEt mixture.
- Product IA is isolated in the form of a yellow syrup (2.73 g, 71%) of E/E and Z/E isomers .
- Example IB Ethyl (2E, 4E) -5 (2-cyano-phenyl) -penta-2, 4-dienoate Compound IA (2.34 g, 10.3 mmol) in solution in acetonitrile (14 ml) is treated at room temperature with iodine (15.0 mg, 0.06 mmol) . After 3 hours of agitation the mixture is evaporated to dryness, taken up in dichloromethane and washed with a Na2SU3 solution (0.01 M) . The organic phase is dried on Na2SO 4 , filtered and evaporated to dryness.
- Product IB is isolated in the form of a solid (2.26 g, 97%) and is used as-is for the following step.
- 1 H NMR, DMSO-d 6 ppm: 1.25 (t, 3H); 4.16 (q, 2H); 6.21 (d, IH); 7.34 (m, 2H); 7.50 (m, 2H); 7.74 (t, IH); 7.87 (d, IH) ; 7.96 (d, IH) .
- Example 1C 5- (2-Cyano-phenyl) -penta-2, 4-dienoic acid Compound IB (2.0 g, 8.82 mmol) in solution in ethanol (50 ml) is treated with 1 N potash (13.2 ml, 13.2 mmol). After 1.5 hours of agitation at reflux, the mixture is evaporated to dryness, taken up with water and treated with 1 N HCl up to acid pH. The precipitate formed is filtered, washed with water and dried under a vacuum to yield pure product 1C (1.63 g, 93%) .
- Acid 1C (700 mg, 3.51 mmol) in solution in dichloromethane (10 ml) in the presence of diisopropylethylamine (DIEA) (1.2 ml, 7.02 mmol) is treated, at room temperature, with 1- (3-dimethylaminopropyl) -3- ethylcarbodiimide (EDCI) hydrochloride (807 mg, 4.21 mmol), 3- hydroxy-1, 2, 3-benzotriazin-4 (3H) -one (HOOBT) (686 mg, 4.21 mmol) and then l-pyridin-2-yl-piperazine (642 ⁇ l, 4.21 mmol) .
- DIEA diisopropylethylamine
- Compound 9 is prepared from intermediate 9B (67.0 mg, 0.32 mmol) and cyclopentylpiperazine (101.3 mg, 0.45 mmol) according to the conditions described for the preparation of compound 1 from 1C. The pure product is isolated in the form of hydrochloride (99 mg, 81%) .
- Example 16A Ethyl 5- (2-nitro-phenyl) -penta-2, 4-dienoate
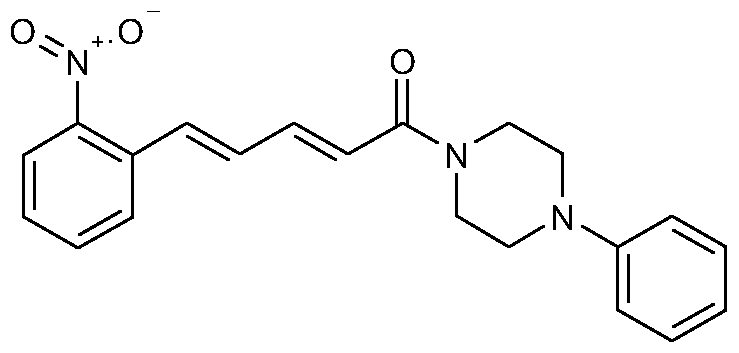
- Example 16 5- (2-Nitro-phenyl) -1- (4-phenyl-piperazin-l-yl) - penta-2, 4-dien-l-one Compound 16 is prepared from intermediate 16B (404 mg,
- Example 27A Ethyl 5- (2, 6-difluoro-phenyl) -penta-2, 4-dienoate Ethyl (diethoxy-phosphoryl) -acetate (3.72 ml, 18.7 mmol) in solution in THF (114 ml) is treated with sodium hydride
- Compound 27 is prepared from intermediate 27B (60 mg, 0.285 mmol) and phenyl-piperazine (68.1 ⁇ l, 0.342 mmol) according to the conditions described for the preparation of compound 1 from 1C.
- the pure product is isolated in the form of beige powder (72 mg, 79%) .
- Example 32A Ethyl 5- (2-fluoro-phenyl) -penta-2, 4-dienoate
- Example 32 1- (4-Cyclopentyl-piperazin-l-yl) -5- (2-fluoro- phenyl) -penta-2, 4-dien-l-one
- Compound 32 is prepared from intermediate 32B (100.0 mg, 0.52 mmol) and cyclopentyl-piperazine (165.3 mg, 0.73 mmol) according to the conditions described for the preparation of compound 1 from 1C. The pure product is isolated in the form of white powder (122 mg, 64%) .
- Mass spectrum (ESI+) m/z 329 (M+H + )
- the derivatives of the present invention are PAR-I receptor antagonists as the results of the models described below demonstrate:
- activation of PAR-I receptors by the SFLLR peptide triggers an intracellular signal cascade leading to the release of calcium by the endoplasmic reticulum.
- Chinese hamster ovarian (CHO) cells constituently express PAR-I receptor.
- the release of calcium consecutive to receptor activation by SFLLR is measured by a fluorometry technique (fluorometric imaging plate reader, or FLIPR) using a selective probe for calcium (Fluo-3AM) .
- FLIPR fluorometric imaging plate reader
- FLIPR selective probe for calcium
- the compounds described in the present invention have demonstrated that they are capable of antagonizing PAR-I receptors and thus decreasing the release of calcium induced by the agonist.
- Materials Culture medium: Ham's F-12 (Ham, R. G., Proc. Nat. Acad. Sci. 1965, 53: 288) supplemented with 10% fetal calf serum and antibiotic (Probenicid, 2.5 mM) .
- Fluorescent probe Fluo-3AM (4 ⁇ M; Teflabs, Austin, Texas, USA)
- Agonist SFLLR-NH2 (Serine, phenylalanine, leucine, leucine, arginine) .
- the derivatives of the present invention were identified as PAR-I receptor antagonists (antagonism >60% of the calcium signal at 10 ⁇ M) .
- the dose-response curves (0.01 ⁇ M to 32 ⁇ M) obtained with the SFLLR agonist allowed determination of the effective concentration inducing 50% of the maximum effect (EC 5 o) .
- the strengths (pA2) of some of the PAR-I antagonists described in the present invention were calculated using the method of Arunlakshana and Schild (Brit. J. Pharmacol., 1959, 14: 48-58) from the EC 50 shifts observed at three concentrations.
- the studies are conducted using guinea pigs (PAR-I receptors similar to man) .
- the carotid flow rate is quantified using a Transonic flow probe. The time required to completely occlude the carotid (flow rate of 0) is measured.
- Methods After the animal is anesthetized (60 mg/kg pentobarbital) , 5 mm of the carotid artery is resected and the laser is placed 4 mm above the artery. A flow probe placed upstream measures occlusion time.
- Rose Bengal (20 mg/kg) is administered by intravenous route and the vessel is irradiated at a wavelength of 514 nm (for 3 min) .
- PAR-I antagonists are administered by intravenous route using a bolus (over 2 min immediately before administration of Rose Bengal) followed by a 15-minute perfusion which begins when the laser is turned on . Results:
- Certain compounds described in the present invention have shown that they are able, after administration by intravenous route at doses from 0.16 mg/kg to 2.5 mg/kg, to delay the time before the formation of a thrombus from 10% to 90% compared to animals receiving vehicle alone.
- the derivatives according to the invention are also of use in the treatment of atrial fibrillation.
- the right and left auricles dilate, thus constituting the substrate for the genesis of atrial fibrillation.
- the disturbance of hemostasis in the cavity of the dilated auricle of a patient suffering from atrial fibrillation leads to an abnormal concentration of thrombin.
- the inventors have demonstrated that this accumulation of thrombin is responsible for an up-regulation of PAR-I which can trigger the proliferation of fibroblasts as well as the formation of platelet thrombus.
- PAR-I antagonists can thus prevent atrial dilation, fibroblast proliferation and thrombus formation in the auricle of a patient suffering from atrial fibrillation .
- a PAR-I antagonist constitutes an effective preventative and/or curative treatment for atrial fibrillation.
- the compounds described in the present invention have demonstrated that they are capable of antagonizing PAR-I receptors and preventing auricle dilation. Materials:
- the animal is anesthetized by a 3.5% mixture of isoflurane in oxygen (Aerrane, Baxter Laboratories) .
- a thoracotomy perpendicular to the sternum of approximately 2 cm is performed at the level of the fourth intercostal space towards the left forefoot.
- a ligature (4-0 silk, CCl needle, Ethicon) is passed around the left coronary artery 1 mm from its origin.
- a surgical knot sufficiently tight to completely occlude the vessel, is tied around the left coronary artery.
- the continuously-recording electrocardiogram makes it possible to verify the satisfactory positioning of the ligature.
- Certain compounds described in the present invention have shown that they are able, after administration by oral route in doses from 10-100 mg/kg/d for 60 days, to reduce by 20% to 90% the auricle surface (measured by echocardiography) compared to untreated animals.
- the present invention also relates to pharmaceutical compositions containing as an active ingredient a compound of general formula (I), or a pharmaceutically-acceptable salt thereof, mixed or combined with a suitable excipient.
- Such compositions can assume the form, for example, of solid or liquid compositions, emulsions, lotions or creams.
- solid compositions for oral administration tablets, pills, powders (in gelatin capsules or in packets) or granules can be used.
- the active ingredient according to the invention is mixed with one or more inert diluents, such as starch, cellulose, sucrose, lactose or silica, under an argon flow.
- Such compositions may also include substances other than diluents, for example one or more lubricants such as magnesium stearate or talc, a colorant, a coating (for sugar-coated pills) or a varnish.
- compositions for oral administration the following can be used: pharmaceutically-acceptable solutions, suspensions, emulsions, syrups and elixirs containing inert diluents such as water, ethanol, glycerol, vegetable oils or liquid paraffin.
- inert diluents such as water, ethanol, glycerol, vegetable oils or liquid paraffin.
- Such compositions can include substances other than diluents, for example wetting, sweetening, thickening, flavoring or stabilizing agents.
- Sterile compositions for parenteral administration can be, preferably, aqueous or non-aqueous solutions, suspensions or emulsions.
- a solvent or vehicle the following can be used: water, propylene glycol, polyethylene glycol, vegetable oils, in particular olive oil, injectable organic esters, for example ethyl oleate or other suitable organic solvents.
- Such compositions can also contain additives, in particular wetting agents, isotonic agents, emulsifiers, dispersants and stabilizers. Sterilization can be achieved in several ways, for example by sterilizing filtration, by incorporating sterilizing agents in the composition, by irradiation or by heating.
- Such compositions can also be prepared in the form of sterile solid compositions that can be dissolved in sterile water or in any other injectable sterile medium just before use .
- compositions for rectal administration are suppositories or rectal capsules that contain, in addition to the active product, excipients such as cocoa butter, semi ⁇ synthetic glycerides or polyethylene glycols.
- compositions for topical administration can be creams, lotions, eye drops, mouth washes, nose drops or aerosols, for example.
- Doses depend on desired effect, treatment duration and administration route, and are generally between 0.001 g and 1 g (preferably between 0.005 g and 0.75 g) per day, preferably by oral route for an adult, with unit doses ranging from 0.1 mg to 500 mg of active substance.
- the doctor will establish suitable dosing according to the patient's age, weight and other specific factors of the case.
- the present invention also relates to products containing a compound according to general formula (I) and another cardiovascular agent as a combination product for simultaneous, separate or time-release use in cardiovascular therapy, the other cardiovascular agent able to be an antiplatelet agent such as aspirin, clopidogrel, ticlopidine, abciximab, tirofiban or eptifibatide .
- an antiplatelet agent such as aspirin, clopidogrel, ticlopidine, abciximab, tirofiban or eptifibatide .
Landscapes
- Chemical & Material Sciences (AREA)
- Organic Chemistry (AREA)
- Health & Medical Sciences (AREA)
- Veterinary Medicine (AREA)
- Animal Behavior & Ethology (AREA)
- Engineering & Computer Science (AREA)
- Public Health (AREA)
- General Health & Medical Sciences (AREA)
- Medicinal Chemistry (AREA)
- Life Sciences & Earth Sciences (AREA)
- Bioinformatics & Cheminformatics (AREA)
- Pharmacology & Pharmacy (AREA)
- Nuclear Medicine, Radiotherapy & Molecular Imaging (AREA)
- General Chemical & Material Sciences (AREA)
- Chemical Kinetics & Catalysis (AREA)
- Cardiology (AREA)
- Heart & Thoracic Surgery (AREA)
- Diabetes (AREA)
- Hematology (AREA)
- Pain & Pain Management (AREA)
- Vascular Medicine (AREA)
- Hospice & Palliative Care (AREA)
- Urology & Nephrology (AREA)
- Rheumatology (AREA)
- Gastroenterology & Hepatology (AREA)
- Pulmonology (AREA)
- Epidemiology (AREA)
- Pharmaceuticals Containing Other Organic And Inorganic Compounds (AREA)
- Heterocyclic Carbon Compounds Containing A Hetero Ring Having Oxygen Or Sulfur (AREA)
- Pyridine Compounds (AREA)
- Heterocyclic Compounds Containing Sulfur Atoms (AREA)
- Thiazole And Isothizaole Compounds (AREA)
Abstract
Description




Claims


Priority Applications (17)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| AU2007263049A AU2007263049B2 (en) | 2006-06-19 | 2007-06-19 | Phenylpentadienoyl derivatives and their use as par 1 antagonists |
| DE602007012888T DE602007012888D1 (en) | 2006-06-19 | 2007-06-19 | PHENYLPENTADIENOYL DERIVATIVES AND THEIR USE AS PAR-1 ANTAGONISTS |
| BRPI0713582-3A BRPI0713582A2 (en) | 2006-06-19 | 2007-06-19 | phenylpentadiene derivatives and their use as antagonists of a pair of |
| JP2009515858A JP5268897B2 (en) | 2006-06-19 | 2007-06-19 | Phenylpentadienoyl derivatives and their use as PAR1 antagonists |
| CN2007800223489A CN101472894B (en) | 2006-06-19 | 2007-06-19 | Phenylpentadiene derivatives and their use as PAR1 antagonists |
| NZ573591A NZ573591A (en) | 2006-06-19 | 2007-06-19 | Phenylpentadienoyl derivatives and their use as par 1 antagonists |
| UAA200900375A UA97370C2 (en) | 2006-06-19 | 2007-06-19 | Phenylpentadienoyl derivatives and their use as par 1 antagonists |
| MX2008016446A MX2008016446A (en) | 2006-06-19 | 2007-06-19 | Phenylpentadienoyl derivatives and their use as par 1 antagonists. |
| EP07765492A EP2046748B1 (en) | 2006-06-19 | 2007-06-19 | Phenylpentadienoyl derivatives and their use as par 1 antagonists |
| US12/305,447 US8022064B2 (en) | 2006-06-19 | 2007-06-19 | Phenylpentadienoyl derivatives and their use as PAR 1 antagonists |
| AT07765492T ATE500223T1 (en) | 2006-06-19 | 2007-06-19 | PHENYLPENTADIENOYL DERIVATIVES AND THEIR USE AS PAR-1 ANTAGONISTS |
| CA002655728A CA2655728A1 (en) | 2006-06-19 | 2007-06-19 | Phenylpentadienoyl derivatives and their use as par 1 antagonists |
| IL195869A IL195869A0 (en) | 2006-06-19 | 2008-12-11 | Phenylpentadienoyl derivatives and their use as par 1 antagonists |
| TNP2008000521A TNSN08521A1 (en) | 2006-06-19 | 2008-12-16 | Phenylpentadienoyl derivatives and their use as par 1 antagonists |
| ZA2008/10629A ZA200810629B (en) | 2006-06-19 | 2008-12-17 | Phenylpentadienoyl derivatives and their use as par 1 antagonists |
| MA31483A MA31131B1 (en) | 2006-06-19 | 2008-12-18 | Phenylpentadienoil derivatives and their use of liabilities. |
| NO20090269A NO20090269L (en) | 2006-06-19 | 2009-01-16 | Phenol pentadienoyl derivatives and their use as PAR-1 antagonists |
Applications Claiming Priority (2)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| FR0605418 | 2006-06-19 | ||
| FR0605418A FR2902427B1 (en) | 2006-06-19 | 2006-06-19 | PHENYLPENTADIENOYLE DERIVATIVES |
Publications (1)
| Publication Number | Publication Date |
|---|---|
| WO2007147822A1 true WO2007147822A1 (en) | 2007-12-27 |
Family
ID=37622010
Family Applications (1)
| Application Number | Title | Priority Date | Filing Date |
|---|---|---|---|
| PCT/EP2007/056078 Ceased WO2007147822A1 (en) | 2006-06-19 | 2007-06-19 | Phenylpentadienoyl derivatives and their use as par 1 antagonists |
Country Status (24)
| Country | Link |
|---|---|
| US (1) | US8022064B2 (en) |
| EP (1) | EP2046748B1 (en) |
| JP (1) | JP5268897B2 (en) |
| KR (1) | KR20090031735A (en) |
| CN (1) | CN101472894B (en) |
| AR (1) | AR061519A1 (en) |
| AT (1) | ATE500223T1 (en) |
| AU (1) | AU2007263049B2 (en) |
| BR (1) | BRPI0713582A2 (en) |
| CA (1) | CA2655728A1 (en) |
| DE (1) | DE602007012888D1 (en) |
| ES (1) | ES2361061T3 (en) |
| FR (1) | FR2902427B1 (en) |
| IL (1) | IL195869A0 (en) |
| MA (1) | MA31131B1 (en) |
| MX (1) | MX2008016446A (en) |
| NO (1) | NO20090269L (en) |
| NZ (1) | NZ573591A (en) |
| RU (1) | RU2440985C2 (en) |
| TN (1) | TNSN08521A1 (en) |
| TW (1) | TW200815371A (en) |
| UA (1) | UA97370C2 (en) |
| WO (1) | WO2007147822A1 (en) |
| ZA (1) | ZA200810629B (en) |
Families Citing this family (3)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| CN104496879A (en) * | 2015-01-13 | 2015-04-08 | 佛山市赛维斯医药科技有限公司 | Nitrile-based benzene and diene fluoro-adamantane contained structure compound and application |
| WO2019190177A1 (en) * | 2018-03-27 | 2019-10-03 | 주식회사 바이오웨이 | (2e,4e)-5-phenyl-penta-2,4-dien-1-one derivative |
| CN118852057B (en) * | 2024-05-29 | 2025-09-26 | 赣南医科大学 | Application of piperine or its derivatives |
Citations (2)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| JPS61106564A (en) * | 1984-10-30 | 1986-05-24 | Terumo Corp | Unsaturated fatty acid amide derivative and inhibitor of blood platelet aggregation containing same |
| JPS61137866A (en) * | 1984-12-07 | 1986-06-25 | Terumo Corp | Imidazole derivative, and platelet coagulation suppressing agent contanining same |
Family Cites Families (3)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| JPS62277370A (en) * | 1986-02-04 | 1987-12-02 | Terumo Corp | Diene derivative and vasodilator containing same |
| EP1818326A1 (en) * | 2000-01-20 | 2007-08-15 | Eisai R&D Management Co., Ltd. | Nitrogen-containing heterocyclic compounds and pharmaceutical composition containing the compounds |
| FR2917622B1 (en) * | 2007-06-19 | 2009-10-23 | Pierre Fabre Medicament Sa | USE OF A PAR1 ANTAGONIST IN THE TREATMENT OF ATRIAL FIBRILLATION. |
-
2006
- 2006-06-19 FR FR0605418A patent/FR2902427B1/en not_active Expired - Fee Related
-
2007
- 2007-06-19 RU RU2008149917/04A patent/RU2440985C2/en not_active IP Right Cessation
- 2007-06-19 ES ES07765492T patent/ES2361061T3/en active Active
- 2007-06-19 AR ARP070102684A patent/AR061519A1/en not_active Application Discontinuation
- 2007-06-19 WO PCT/EP2007/056078 patent/WO2007147822A1/en not_active Ceased
- 2007-06-19 JP JP2009515858A patent/JP5268897B2/en not_active Expired - Fee Related
- 2007-06-19 KR KR1020097000945A patent/KR20090031735A/en not_active Ceased
- 2007-06-19 US US12/305,447 patent/US8022064B2/en not_active Expired - Fee Related
- 2007-06-19 DE DE602007012888T patent/DE602007012888D1/en active Active
- 2007-06-19 AT AT07765492T patent/ATE500223T1/en not_active IP Right Cessation
- 2007-06-19 CN CN2007800223489A patent/CN101472894B/en not_active Expired - Fee Related
- 2007-06-19 NZ NZ573591A patent/NZ573591A/en not_active IP Right Cessation
- 2007-06-19 EP EP07765492A patent/EP2046748B1/en active Active
- 2007-06-19 UA UAA200900375A patent/UA97370C2/en unknown
- 2007-06-19 CA CA002655728A patent/CA2655728A1/en not_active Abandoned
- 2007-06-19 BR BRPI0713582-3A patent/BRPI0713582A2/en not_active Application Discontinuation
- 2007-06-19 AU AU2007263049A patent/AU2007263049B2/en not_active Ceased
- 2007-06-19 MX MX2008016446A patent/MX2008016446A/en active IP Right Grant
- 2007-06-20 TW TW096122075A patent/TW200815371A/en unknown
-
2008
- 2008-12-11 IL IL195869A patent/IL195869A0/en unknown
- 2008-12-16 TN TNP2008000521A patent/TNSN08521A1/en unknown
- 2008-12-17 ZA ZA2008/10629A patent/ZA200810629B/en unknown
- 2008-12-18 MA MA31483A patent/MA31131B1/en unknown
-
2009
- 2009-01-16 NO NO20090269A patent/NO20090269L/en not_active Application Discontinuation
Patent Citations (2)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| JPS61106564A (en) * | 1984-10-30 | 1986-05-24 | Terumo Corp | Unsaturated fatty acid amide derivative and inhibitor of blood platelet aggregation containing same |
| JPS61137866A (en) * | 1984-12-07 | 1986-06-25 | Terumo Corp | Imidazole derivative, and platelet coagulation suppressing agent contanining same |
Non-Patent Citations (3)
| Title |
|---|
| CHACKALAMANNIL S ET AL: "Thrombin receptor (PAR-1) antagonists as novel antithrombotic agents", EXPERT OPINION ON THERAPEUTIC PATENTS 2006 UNITED KINGDOM, vol. 16, no. 4, April 2006 (2006-04-01), pages 493 - 505, XP002415653, ISSN: 1354-3776 * |
| DATABASE WPI Week 198627, Derwent World Patents Index; AN 1986-173349, XP002416150 * |
| DATABASE WPI Week 198632, Derwent World Patents Index; AN 1986-207178, XP002416149 * |
Also Published As
| Publication number | Publication date |
|---|---|
| RU2440985C2 (en) | 2012-01-27 |
| US8022064B2 (en) | 2011-09-20 |
| MX2008016446A (en) | 2009-01-22 |
| US20100003260A1 (en) | 2010-01-07 |
| JP5268897B2 (en) | 2013-08-21 |
| MA31131B1 (en) | 2010-02-01 |
| IL195869A0 (en) | 2009-09-01 |
| CN101472894A (en) | 2009-07-01 |
| NZ573591A (en) | 2010-10-29 |
| TNSN08521A1 (en) | 2010-04-14 |
| ZA200810629B (en) | 2010-02-24 |
| NO20090269L (en) | 2009-02-27 |
| UA97370C2 (en) | 2012-02-10 |
| JP2009541258A (en) | 2009-11-26 |
| RU2008149917A (en) | 2010-07-27 |
| ATE500223T1 (en) | 2011-03-15 |
| DE602007012888D1 (en) | 2011-04-14 |
| TW200815371A (en) | 2008-04-01 |
| BRPI0713582A2 (en) | 2012-10-23 |
| CN101472894B (en) | 2012-09-05 |
| AR061519A1 (en) | 2008-09-03 |
| CA2655728A1 (en) | 2007-12-27 |
| KR20090031735A (en) | 2009-03-27 |
| FR2902427B1 (en) | 2008-08-22 |
| EP2046748B1 (en) | 2011-03-02 |
| AU2007263049B2 (en) | 2011-10-27 |
| FR2902427A1 (en) | 2007-12-21 |
| AU2007263049A1 (en) | 2007-12-27 |
| EP2046748A1 (en) | 2009-04-15 |
| ES2361061T3 (en) | 2011-06-13 |
Similar Documents
| Publication | Publication Date | Title |
|---|---|---|
| EP2041104B1 (en) | Cinnamoyl-piperazine derivatives and their use as par- i antagonists | |
| EP2046748B1 (en) | Phenylpentadienoyl derivatives and their use as par 1 antagonists | |
| HK1129385A (en) | Phenylpentadienoyl derivatives and their use as par 1 antagonists | |
| HK1127353B (en) | Cinnamoyl-piperazine derivatives and their use as par- i antagonists | |
| WO2006043510A1 (en) | Ester derivative and pharmaceutical use thereof |
Legal Events
| Date | Code | Title | Description |
|---|---|---|---|
| WWE | Wipo information: entry into national phase |
Ref document number: 200780022348.9 Country of ref document: CN |
|
| 121 | Ep: the epo has been informed by wipo that ep was designated in this application |
Ref document number: 07765492 Country of ref document: EP Kind code of ref document: A1 |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 10233/DELNP/2008 Country of ref document: IN |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 2007263049 Country of ref document: AU Ref document number: 195869 Country of ref document: IL |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 573591 Country of ref document: NZ |
|
| ENP | Entry into the national phase |
Ref document number: 2655728 Country of ref document: CA |
|
| WWE | Wipo information: entry into national phase |
Ref document number: DZP2008000729 Country of ref document: DZ |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 12305447 Country of ref document: US Ref document number: MX/A/2008/016446 Country of ref document: MX |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 2009515858 Country of ref document: JP |
|
| NENP | Non-entry into the national phase |
Ref country code: DE |
|
| ENP | Entry into the national phase |
Ref document number: 2007263049 Country of ref document: AU Date of ref document: 20070619 Kind code of ref document: A |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 1020097000945 Country of ref document: KR Ref document number: 2007765492 Country of ref document: EP |
|
| ENP | Entry into the national phase |
Ref document number: 2008149917 Country of ref document: RU Kind code of ref document: A |
|
| ENP | Entry into the national phase |
Ref document number: PI0713582 Country of ref document: BR Kind code of ref document: A2 Effective date: 20081218 |