WO2013127441A1 - Process for the preparation of substituted phenylpropanones - Google Patents
Process for the preparation of substituted phenylpropanones Download PDFInfo
- Publication number
- WO2013127441A1 WO2013127441A1 PCT/EP2012/053371 EP2012053371W WO2013127441A1 WO 2013127441 A1 WO2013127441 A1 WO 2013127441A1 EP 2012053371 W EP2012053371 W EP 2012053371W WO 2013127441 A1 WO2013127441 A1 WO 2013127441A1
- Authority
- WO
- WIPO (PCT)
- Prior art keywords
- formula
- compound
- preparation
- crc
- alkyl
- Prior art date
- Legal status (The legal status is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the status listed.)
- Ceased
Links
- 0 Cc1cc(*)cc(*)c1N Chemical compound Cc1cc(*)cc(*)c1N 0.000 description 1
Classifications
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07C—ACYCLIC OR CARBOCYCLIC COMPOUNDS
- C07C45/00—Preparation of compounds having >C = O groups bound only to carbon or hydrogen atoms; Preparation of chelates of such compounds
- C07C45/51—Preparation of compounds having >C = O groups bound only to carbon or hydrogen atoms; Preparation of chelates of such compounds by pyrolysis, rearrangement or decomposition
- C07C45/516—Preparation of compounds having >C = O groups bound only to carbon or hydrogen atoms; Preparation of chelates of such compounds by pyrolysis, rearrangement or decomposition involving transformation of nitrogen-containing compounds to >C = O groups
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07C—ACYCLIC OR CARBOCYCLIC COMPOUNDS
- C07C45/00—Preparation of compounds having >C = O groups bound only to carbon or hydrogen atoms; Preparation of chelates of such compounds
- C07C45/45—Preparation of compounds having >C = O groups bound only to carbon or hydrogen atoms; Preparation of chelates of such compounds by condensation
- C07C45/455—Preparation of compounds having >C = O groups bound only to carbon or hydrogen atoms; Preparation of chelates of such compounds by condensation with carboxylic acids or their derivatives
Definitions
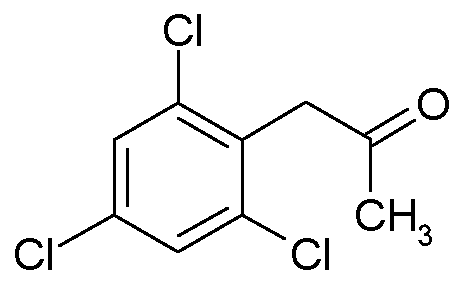
- the present invention relates to a process for the preparation of propane-2-one-substituted phenyls, in particular to the preparation of 1-(2,4,6-trihalo-phenyl)-propan-2-ones.
- Propane-2-one-substituted phenyls are valuable intermediates for the preparation of fungicidally active pyrazole carboxamides, as described for example in
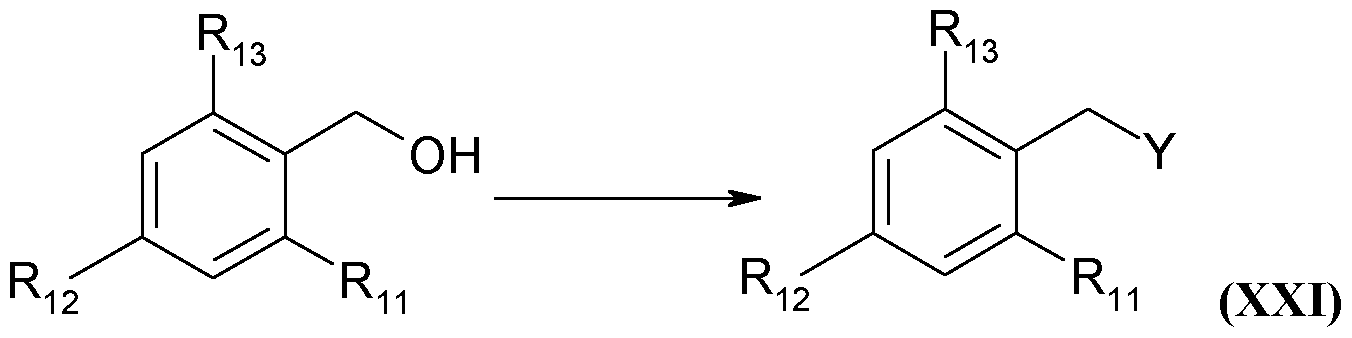
- Rn is hydrogen, halogen or d-C 6 alkyl
- R 12 is hydrogen, halogen, CrC 6 alkyl, C 2 -C 6 alkenyl, C 3 -C 6 alkinyl, C3-C 6 cycloalkyl-C 3 - C 6 alkinyl, halophenoxy, halophenyl-C 3 -C 6 alkinyl, C
- R 13 is hydrogen, halogen or CrC 6 alkyl
- a disadvantage of this prior art process is the high production costs caused by the significant number of reaction steps which makes this process uneconomical and especially unsuitable for a large-scale production.
- WO 00/34229 describes a process for the preparation of a ketone of formula V
- Diazonium salts in general are sensitive to physical agents such as heat, light, shock, static electricity and dehydration that can lead to rapid, uncontrollable decompositions and explosions.
- a further disadvantage is that 2 different equipments are needed to perform the reactions.
- the aim of the present invention is therefore to provide a novel process for the production of
- propane-2-one-substituted phenyls that avoids the disadvantages of the known process and makes it possible to prepare propane-2-one-substituted phenyls in high yields and good quality in an economically advantageous way with less reaction steps.
- R is hydrogen, halogen or CrC 6 alkyl
- R 2 is hydrogen, halogen, CrC 6 alkyl, C 2 -C 6 alkenyl, C 3 -C 6 alkinyl, C 3 -C 6 cycloalkyl-C 3 -C 6 alkinyl, halophenoxy, halophenyl-C 3 -C 6 alkinyl, CrC 6 haloalkyl, CrC 6 haloalkoxy, C 2 -C 6 haloalkenyl or C 2 -C 6 haloalkenyloxy; and R 3 is hydrogen, halogen or d-C 6 alkyl;
- R 2 and R 3 have the meanings as described under formula I, in the presence of an inert organic solvent, to a mixture comprising an organic nitrite of formula III
- R 4 -0-N 0 (III), wherein R 4 is CrC 8 alkyl, a compound of formula IV
- the process according to the invention uses easily accessible and nontoxic starting material, without the need of isolation or accumulation of diazonium salt and is therefore especially suitable for the large-scale preparation of a compound of formula I.
- the process according to the invention is especially suitable for the production of compounds of formula I, wherein at least one of R ⁇ R 2 and R 3 is different from hydrogen.
- the process according to the invention is especially suitable for the preparation of compounds of formula I, wherein at least one of R ⁇ R 2 and R 3 is halogen.
- the compound of formula V can be prepared preferably by a one pot reaction adding the aniline of formula II to a mixture of isoprenylacetate of formula IV, an organic nitrite of formula III and a solvent.
- R 4 is C 4 -C 7 alkyl.
- a preferred nitrite is tert-butyl nitrite.
- the mixture of isoprenylacetate of formula IV, an organic nitrite of formula III and a solvent can additionally contain a copper compound which can be advantageous to increase yield and/or quality of the product.
- Preferred copper compounds are CuO, CuCI 2 or CuSCv
- the amount of the copper compounds is preferably is 1-20 mol% in the relation to the aniline of formula II.
- the reaction is a temperature of -10°C to 50°C. No isolation or accumulation of the diazonium salt is required for this process step.
- the same solvent is used for the aniline of formula II and the mixture of isoprenylacetate of formula IV, and the organic nitrite of formula III.
- Suitable inert organic solvents are for example ketones, for example acetone, methylethylketone (MEK) or nitriles, for example acetonitrile.
- Preferred solvents are acetone and acetonitrile.
- the compounds of formula II and III are either known or can be prepared according to methods well known in the art. Some compounds of formula II are commercially available, e.g. the compound of formula II, wherein R 2 and R 3 is chloro. Isoprenylacetate of formula IV is commercially available.
Landscapes
- Chemical & Material Sciences (AREA)
- Organic Chemistry (AREA)
- Organic Low-Molecular-Weight Compounds And Preparation Thereof (AREA)
Abstract
Description






Claims


Priority Applications (17)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| PT127081057T PT2819983T (en) | 2012-02-28 | 2012-02-28 | Process for the preparation of substituted phenylpropanones |
| MX2014009829A MX349740B (en) | 2012-02-28 | 2012-02-28 | Process for the preparation of substituted phenylpropanones. |
| PL12708105T PL2819983T3 (en) | 2012-02-28 | 2012-02-28 | Process for the preparation of substituted phenylpropanones |
| PCT/EP2012/053371 WO2013127441A1 (en) | 2012-02-28 | 2012-02-28 | Process for the preparation of substituted phenylpropanones |
| KR1020147024525A KR101902373B1 (en) | 2012-02-28 | 2012-02-28 | Process for the preparation of substituted phenylpropanones |
| ES12708105.7T ES2652365T3 (en) | 2012-02-28 | 2012-02-28 | Process for the preparation of substituted phenylpropanones |
| JP2014558015A JP6251197B2 (en) | 2012-02-28 | 2012-02-28 | Process for preparing substituted phenylpropanones |
| BR112014021063-2A BR112014021063B1 (en) | 2012-02-28 | 2012-02-28 | PROCESS FOR THE PREPARATION OF REPLACED PHENYLPROPANONES |
| HUE12708105A HUE037597T2 (en) | 2012-02-28 | 2012-02-28 | Process for the preparation of substituted phenylpropanones |
| CA2863420A CA2863420C (en) | 2012-02-28 | 2012-02-28 | Process for the preparation of substituted phenylpropanones |
| EP12708105.7A EP2819983B1 (en) | 2012-02-28 | 2012-02-28 | Process for the preparation of substituted phenylpropanones |
| CN201280070805.2A CN104136409B (en) | 2012-02-28 | 2012-02-28 | Process for the preparation of substituted propiophenones |
| US14/379,013 US9073839B2 (en) | 2012-02-28 | 2012-02-28 | Process for the preparation of substituted phenylpropanones |
| DK12708105.7T DK2819983T3 (en) | 2012-02-28 | 2012-02-28 | Process for preparing substituted phenylpropanones |
| AU2012371236A AU2012371236B2 (en) | 2012-02-28 | 2012-02-28 | Process for the preparation of substituted phenylpropanones |
| IL233938A IL233938A (en) | 2012-02-28 | 2014-08-04 | Process for the preparation of substituted phenylpropanones |
| ZA2014/05824A ZA201405824B (en) | 2012-02-28 | 2014-08-07 | Process for the preparation of substituted phenylpropanones |
Applications Claiming Priority (1)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| PCT/EP2012/053371 WO2013127441A1 (en) | 2012-02-28 | 2012-02-28 | Process for the preparation of substituted phenylpropanones |
Publications (1)
| Publication Number | Publication Date |
|---|---|
| WO2013127441A1 true WO2013127441A1 (en) | 2013-09-06 |
Family
ID=45814485
Family Applications (1)
| Application Number | Title | Priority Date | Filing Date |
|---|---|---|---|
| PCT/EP2012/053371 Ceased WO2013127441A1 (en) | 2012-02-28 | 2012-02-28 | Process for the preparation of substituted phenylpropanones |
Country Status (17)
| Country | Link |
|---|---|
| US (1) | US9073839B2 (en) |
| EP (1) | EP2819983B1 (en) |
| JP (1) | JP6251197B2 (en) |
| KR (1) | KR101902373B1 (en) |
| CN (1) | CN104136409B (en) |
| AU (1) | AU2012371236B2 (en) |
| BR (1) | BR112014021063B1 (en) |
| CA (1) | CA2863420C (en) |
| DK (1) | DK2819983T3 (en) |
| ES (1) | ES2652365T3 (en) |
| HU (1) | HUE037597T2 (en) |
| IL (1) | IL233938A (en) |
| MX (1) | MX349740B (en) |
| PL (1) | PL2819983T3 (en) |
| PT (1) | PT2819983T (en) |
| WO (1) | WO2013127441A1 (en) |
| ZA (1) | ZA201405824B (en) |
Cited By (2)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| JP2024056863A (en) * | 2018-09-06 | 2024-04-23 | エフ エム シー コーポレーション | Fungicidal nitroanilino-substituted pyrazoles. |
| WO2026131414A1 (en) * | 2024-12-18 | 2026-06-25 | Syngenta Crop Protection Ag | Chemical process for the production of pydiflumetofen |
Families Citing this family (3)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| CN108610290B (en) * | 2018-06-11 | 2020-04-24 | 武汉工程大学 | Preparation method of fluxapyroxad |
| CN113004131B (en) * | 2021-02-01 | 2022-05-13 | 江苏先导药业有限公司 | Preparation method of 2,4, 6-trichlorophenyl substituted acetone |
| CN115703701B (en) * | 2021-08-16 | 2024-06-25 | 上海晓明检测技术服务有限公司 | Method for continuously synthesizing 1- (2, 4, 6-trichloro-phenyl) -propan-2-one by micro-channel |
Citations (3)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| WO2000034229A1 (en) | 1998-12-10 | 2000-06-15 | Bayer Aktiengesellschaft | Process for the preparation of strobilurin intermediates |
| US20080221198A1 (en) * | 2005-03-10 | 2008-09-11 | Dalian Zhao | Novel Crystalline Forms of Antidiabetic Compounds |
| WO2010063700A2 (en) | 2008-12-05 | 2010-06-10 | Syngenta Participations Ag | Novel microbiocides |
Family Cites Families (1)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| JP2013100235A (en) * | 2010-03-08 | 2013-05-23 | Dainippon Sumitomo Pharma Co Ltd | Novel indole derivative |
-
2012
- 2012-02-28 HU HUE12708105A patent/HUE037597T2/en unknown
- 2012-02-28 MX MX2014009829A patent/MX349740B/en active IP Right Grant
- 2012-02-28 ES ES12708105.7T patent/ES2652365T3/en active Active
- 2012-02-28 PT PT127081057T patent/PT2819983T/en unknown
- 2012-02-28 BR BR112014021063-2A patent/BR112014021063B1/en active IP Right Grant
- 2012-02-28 CN CN201280070805.2A patent/CN104136409B/en active Active
- 2012-02-28 PL PL12708105T patent/PL2819983T3/en unknown
- 2012-02-28 WO PCT/EP2012/053371 patent/WO2013127441A1/en not_active Ceased
- 2012-02-28 CA CA2863420A patent/CA2863420C/en active Active
- 2012-02-28 DK DK12708105.7T patent/DK2819983T3/en active
- 2012-02-28 US US14/379,013 patent/US9073839B2/en active Active
- 2012-02-28 KR KR1020147024525A patent/KR101902373B1/en active Active
- 2012-02-28 JP JP2014558015A patent/JP6251197B2/en active Active
- 2012-02-28 AU AU2012371236A patent/AU2012371236B2/en active Active
- 2012-02-28 EP EP12708105.7A patent/EP2819983B1/en active Active
-
2014
- 2014-08-04 IL IL233938A patent/IL233938A/en active IP Right Grant
- 2014-08-07 ZA ZA2014/05824A patent/ZA201405824B/en unknown
Patent Citations (3)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| WO2000034229A1 (en) | 1998-12-10 | 2000-06-15 | Bayer Aktiengesellschaft | Process for the preparation of strobilurin intermediates |
| US20080221198A1 (en) * | 2005-03-10 | 2008-09-11 | Dalian Zhao | Novel Crystalline Forms of Antidiabetic Compounds |
| WO2010063700A2 (en) | 2008-12-05 | 2010-06-10 | Syngenta Participations Ag | Novel microbiocides |
Non-Patent Citations (1)
| Title |
|---|
| LI LIA ET AL: "Facile Synthesis of 1-Aryl-2-propanones from Aromatic Amine", SYNTHETIC COMMUNICATIONS,, vol. 37, no. 6, 31 December 2007 (2007-12-31), pages 985 - 991, XP002666726, ISSN: 0039-7911, [retrieved on 20070224], DOI: 10.1080/00397910601163950 * |
Cited By (4)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| JP2024056863A (en) * | 2018-09-06 | 2024-04-23 | エフ エム シー コーポレーション | Fungicidal nitroanilino-substituted pyrazoles. |
| EP4388876A3 (en) * | 2018-09-06 | 2025-05-14 | FMC Corporation | Fungicidal nitroanilino substituted pyrazoles |
| JP7695421B2 (en) | 2018-09-06 | 2025-06-18 | エフ エム シー コーポレーション | Fungicidal nitroanilino-substituted pyrazoles. |
| WO2026131414A1 (en) * | 2024-12-18 | 2026-06-25 | Syngenta Crop Protection Ag | Chemical process for the production of pydiflumetofen |
Also Published As
| Publication number | Publication date |
|---|---|
| KR101902373B1 (en) | 2018-09-28 |
| AU2012371236B2 (en) | 2017-01-05 |
| HUE037597T2 (en) | 2018-09-28 |
| IL233938A0 (en) | 2014-09-30 |
| AU2012371236A1 (en) | 2014-09-18 |
| PL2819983T3 (en) | 2018-02-28 |
| DK2819983T3 (en) | 2018-01-02 |
| ES2652365T3 (en) | 2018-02-01 |
| EP2819983A1 (en) | 2015-01-07 |
| BR112014021063B1 (en) | 2019-08-20 |
| IL233938A (en) | 2017-01-31 |
| EP2819983B1 (en) | 2017-10-04 |
| JP2015511578A (en) | 2015-04-20 |
| JP6251197B2 (en) | 2017-12-20 |
| MX349740B (en) | 2017-08-10 |
| US9073839B2 (en) | 2015-07-07 |
| CN104136409A (en) | 2014-11-05 |
| CA2863420C (en) | 2019-02-19 |
| PT2819983T (en) | 2017-12-18 |
| MX2014009829A (en) | 2014-09-11 |
| US20150031919A1 (en) | 2015-01-29 |
| CN104136409B (en) | 2016-05-11 |
| KR20140131527A (en) | 2014-11-13 |
| CA2863420A1 (en) | 2013-09-06 |
| ZA201405824B (en) | 2015-11-25 |
Similar Documents
| Publication | Publication Date | Title |
|---|---|---|
| CA2863467C (en) | Process for the preparation of phenyl substituted 3-difluoromethyl-1-methyl-1h-pyrazole-4-carboxylic acid n-methoxy- [1-methyl-2-phenylethyl] amides | |
| EP2230234B1 (en) | Process for the preparation of rufinamide | |
| EP2819983B1 (en) | Process for the preparation of substituted phenylpropanones | |
| WO2011026937A1 (en) | Process for preparing 1-phenylpyrazoles | |
| JP5512551B2 (en) | Process for preparing substituted biphenylanilides | |
| CN110740995B (en) | Process for preparing substituted phenoxyphenyl alcohols | |
| EP2980079A1 (en) | Method of manufacturing pyridazinone compound | |
| CA2587554A1 (en) | Process for the production of anilines | |
| WO2013037291A1 (en) | Process for the preparation of n-substituted pyrazole compounds | |
| CN120858093A (en) | Preparation of herbicidal uracil compounds | |
| CN105251497A (en) | Solid catalyst, and preparation method and applications thereof | |
| JP5142241B2 (en) | Method for producing nicotinic acid ester compound | |
| MX2008010140A (en) | Improved method for producing nitroisourea derivative |
Legal Events
| Date | Code | Title | Description |
|---|---|---|---|
| WWE | Wipo information: entry into national phase |
Ref document number: 201280070805.2 Country of ref document: CN |
|
| 121 | Ep: the epo has been informed by wipo that ep was designated in this application |
Ref document number: 12708105 Country of ref document: EP Kind code of ref document: A1 |
|
| ENP | Entry into the national phase |
Ref document number: 2863420 Country of ref document: CA |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 233938 Country of ref document: IL |
|
| WWE | Wipo information: entry into national phase |
Ref document number: MX/A/2014/009829 Country of ref document: MX |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 14379013 Country of ref document: US |
|
| ENP | Entry into the national phase |
Ref document number: 2014558015 Country of ref document: JP Kind code of ref document: A |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 14188185 Country of ref document: CO |
|
| NENP | Non-entry into the national phase |
Ref country code: DE |
|
| ENP | Entry into the national phase |
Ref document number: 20147024525 Country of ref document: KR Kind code of ref document: A |
|
| REEP | Request for entry into the european phase |
Ref document number: 2012708105 Country of ref document: EP |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 2012708105 Country of ref document: EP |
|
| ENP | Entry into the national phase |
Ref document number: 2012371236 Country of ref document: AU Date of ref document: 20120228 Kind code of ref document: A |
|
| REG | Reference to national code |
Ref country code: BR Ref legal event code: B01A Ref document number: 112014021063 Country of ref document: BR |
|
| ENP | Entry into the national phase |
Ref document number: 112014021063 Country of ref document: BR Kind code of ref document: A2 Effective date: 20140826 |