WO2019204251A1 - Therapeutic methods and compositions for treating prostate cancer using 6,8-bis-benzylthio-octanoic acid - Google Patents
Therapeutic methods and compositions for treating prostate cancer using 6,8-bis-benzylthio-octanoic acid Download PDFInfo
- Publication number
- WO2019204251A1 WO2019204251A1 PCT/US2019/027597 US2019027597W WO2019204251A1 WO 2019204251 A1 WO2019204251 A1 WO 2019204251A1 US 2019027597 W US2019027597 W US 2019027597W WO 2019204251 A1 WO2019204251 A1 WO 2019204251A1
- Authority
- WO
- WIPO (PCT)
- Prior art keywords
- therapeutic agent
- administered
- prostate cancer
- benzylthio
- bis
- Prior art date
- Legal status (The legal status is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the status listed.)
- Ceased
Links
Classifications
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61K—PREPARATIONS FOR MEDICAL, DENTAL OR TOILETRY PURPOSES
- A61K31/00—Medicinal preparations containing organic active ingredients
- A61K31/185—Acids; Anhydrides, halides or salts thereof, e.g. sulfur acids, imidic, hydrazonic or hydroximic acids
- A61K31/19—Carboxylic acids, e.g. valproic acid
- A61K31/20—Carboxylic acids, e.g. valproic acid having a carboxyl group bound to a chain of seven or more carbon atoms, e.g. stearic, palmitic, arachidic acids
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61K—PREPARATIONS FOR MEDICAL, DENTAL OR TOILETRY PURPOSES
- A61K31/00—Medicinal preparations containing organic active ingredients
- A61K31/13—Amines
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61K—PREPARATIONS FOR MEDICAL, DENTAL OR TOILETRY PURPOSES
- A61K31/00—Medicinal preparations containing organic active ingredients
- A61K31/13—Amines
- A61K31/133—Amines having hydroxy groups, e.g. sphingosine
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61K—PREPARATIONS FOR MEDICAL, DENTAL OR TOILETRY PURPOSES
- A61K31/00—Medicinal preparations containing organic active ingredients
- A61K31/33—Heterocyclic compounds
- A61K31/335—Heterocyclic compounds having oxygen as the only ring hetero atom, e.g. fungichromin
- A61K31/337—Heterocyclic compounds having oxygen as the only ring hetero atom, e.g. fungichromin having four-membered rings, e.g. taxol
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61K—PREPARATIONS FOR MEDICAL, DENTAL OR TOILETRY PURPOSES
- A61K47/00—Medicinal preparations characterised by the non-active ingredients used, e.g. carriers or inert additives; Targeting or modifying agents chemically bound to the active ingredient
- A61K47/06—Organic compounds, e.g. natural or synthetic hydrocarbons, polyolefins, mineral oil, petrolatum or ozokerite
- A61K47/08—Organic compounds, e.g. natural or synthetic hydrocarbons, polyolefins, mineral oil, petrolatum or ozokerite containing oxygen, e.g. ethers, acetals, ketones, quinones, aldehydes, peroxides
- A61K47/14—Esters of carboxylic acids, e.g. fatty acid monoglycerides, medium-chain triglycerides, parabens or PEG fatty acid esters
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61K—PREPARATIONS FOR MEDICAL, DENTAL OR TOILETRY PURPOSES
- A61K47/00—Medicinal preparations characterised by the non-active ingredients used, e.g. carriers or inert additives; Targeting or modifying agents chemically bound to the active ingredient
- A61K47/06—Organic compounds, e.g. natural or synthetic hydrocarbons, polyolefins, mineral oil, petrolatum or ozokerite
- A61K47/16—Organic compounds, e.g. natural or synthetic hydrocarbons, polyolefins, mineral oil, petrolatum or ozokerite containing nitrogen, e.g. nitro-, nitroso-, azo-compounds, nitriles, cyanates
- A61K47/18—Amines; Amides; Ureas; Quaternary ammonium compounds; Amino acids; Oligopeptides having up to five amino acids
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61K—PREPARATIONS FOR MEDICAL, DENTAL OR TOILETRY PURPOSES
- A61K47/00—Medicinal preparations characterised by the non-active ingredients used, e.g. carriers or inert additives; Targeting or modifying agents chemically bound to the active ingredient
- A61K47/06—Organic compounds, e.g. natural or synthetic hydrocarbons, polyolefins, mineral oil, petrolatum or ozokerite
- A61K47/26—Carbohydrates, e.g. sugar alcohols, amino sugars, nucleic acids, mono-, di- or oligo-saccharides; Derivatives thereof, e.g. polysorbates, sorbitan fatty acid esters or glycyrrhizin
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61K—PREPARATIONS FOR MEDICAL, DENTAL OR TOILETRY PURPOSES
- A61K47/00—Medicinal preparations characterised by the non-active ingredients used, e.g. carriers or inert additives; Targeting or modifying agents chemically bound to the active ingredient
- A61K47/30—Macromolecular organic or inorganic compounds, e.g. inorganic polyphosphates
- A61K47/32—Macromolecular compounds obtained by reactions only involving carbon-to-carbon unsaturated bonds, e.g. carbomers, poly(meth)acrylates, or polyvinyl pyrrolidone
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61K—PREPARATIONS FOR MEDICAL, DENTAL OR TOILETRY PURPOSES
- A61K47/00—Medicinal preparations characterised by the non-active ingredients used, e.g. carriers or inert additives; Targeting or modifying agents chemically bound to the active ingredient
- A61K47/30—Macromolecular organic or inorganic compounds, e.g. inorganic polyphosphates
- A61K47/36—Polysaccharides; Derivatives thereof, e.g. gums, starch, alginate, dextrin, hyaluronic acid, chitosan, inulin, agar or pectin
- A61K47/38—Cellulose; Derivatives thereof
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61K—PREPARATIONS FOR MEDICAL, DENTAL OR TOILETRY PURPOSES
- A61K9/00—Medicinal preparations characterised by special physical form
- A61K9/0012—Galenical forms characterised by the site of application
- A61K9/0019—Injectable compositions; Intramuscular, intravenous, arterial, subcutaneous administration; Compositions to be administered through the skin in an invasive manner
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61K—PREPARATIONS FOR MEDICAL, DENTAL OR TOILETRY PURPOSES
- A61K9/00—Medicinal preparations characterised by special physical form
- A61K9/0012—Galenical forms characterised by the site of application
- A61K9/0053—Mouth and digestive tract, i.e. intraoral and peroral administration
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61K—PREPARATIONS FOR MEDICAL, DENTAL OR TOILETRY PURPOSES
- A61K9/00—Medicinal preparations characterised by special physical form
- A61K9/08—Solutions
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61K—PREPARATIONS FOR MEDICAL, DENTAL OR TOILETRY PURPOSES
- A61K9/00—Medicinal preparations characterised by special physical form
- A61K9/10—Dispersions; Emulsions
- A61K9/107—Emulsions ; Emulsion preconcentrates; Micelles
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61K—PREPARATIONS FOR MEDICAL, DENTAL OR TOILETRY PURPOSES
- A61K9/00—Medicinal preparations characterised by special physical form
- A61K9/14—Particulate form, e.g. powders, Processes for size reducing of pure drugs or the resulting products, Pure drug nanoparticles
- A61K9/16—Agglomerates; Granulates; Microbeadlets ; Microspheres; Pellets; Solid products obtained by spray drying, spray freeze drying, spray congealing,(multiple) emulsion solvent evaporation or extraction
- A61K9/1605—Excipients; Inactive ingredients
- A61K9/1629—Organic macromolecular compounds
- A61K9/1635—Organic macromolecular compounds obtained by reactions only involving carbon-to-carbon unsaturated bonds, e.g. polyvinyl pyrrolidone, poly(meth)acrylates
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61K—PREPARATIONS FOR MEDICAL, DENTAL OR TOILETRY PURPOSES
- A61K9/00—Medicinal preparations characterised by special physical form
- A61K9/14—Particulate form, e.g. powders, Processes for size reducing of pure drugs or the resulting products, Pure drug nanoparticles
- A61K9/16—Agglomerates; Granulates; Microbeadlets ; Microspheres; Pellets; Solid products obtained by spray drying, spray freeze drying, spray congealing,(multiple) emulsion solvent evaporation or extraction
- A61K9/1605—Excipients; Inactive ingredients
- A61K9/1629—Organic macromolecular compounds
- A61K9/1652—Polysaccharides, e.g. alginate, cellulose derivatives; Cyclodextrin
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P35/00—Antineoplastic agents
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P35/00—Antineoplastic agents
- A61P35/04—Antineoplastic agents specific for metastasis
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61K—PREPARATIONS FOR MEDICAL, DENTAL OR TOILETRY PURPOSES
- A61K2300/00—Mixtures or combinations of active ingredients, wherein at least one active ingredient is fully defined in groups A61K31/00 - A61K41/00
Definitions
- the invention provides methods, compositions, and medical kits for treating prostate cancer using 6,8-bis-benzylthio-octanoic acid (CPI-613) or a pharmaceutically acceptable salt thereof, including combination therapy with docetaxel.
- CPI-613 6,8-bis-benzylthio-octanoic acid
- a pharmaceutically acceptable salt thereof including combination therapy with docetaxel.
- Cancer is a leading cause of death in many industrialized countries. Recent estimates are that 10 million Americans are currently living with cancer, and that 1.2 million Americans are newly diagnosed with cancer each year. Significant advances have been made in improving the diagnosis and treatment of cancer. However, current treatment options often suffer from severe adverse side effects and/or the treatments are not effective for all patients. For example, many clinically-accepted chemotherapeutic agents can induce profound damage to normal, proliferative host cells. Another problem associated with many chemotherapeutic treatments is that, in many tumor types, there is either inherent or acquired resistance to the therapy.
- Prostate cancer affects a substantial portion of the male patient population.
- Prostate cancer is characterized by the formation of malignant cells in the prostate.
- Exemplary therapies currently used to help treat patients suffering from prostate cancer include surgery, radiation therapy, and chemotherapy.
- Surgery often includes the complete surgical removal of the prostate gland.
- Radiation therapy involves applying ionizing radiation to the diseased area of the prostate.
- the invention provides methods, compositions, and medical kits for treating prostate cancer using 6,8-bis-benzylthio-octanoic acid or a pharmaceutically acceptable salt thereof, including combination therapy with docetaxel.
- the prostate cancer may be, for example, characterized as an androgen-dependent prostate cancer or an androgen-independent prostate cancer.
- the 6,8-bis-benzylthio-octanoic acid or a pharmaceutically acceptable salt thereof may be formulated as a pharmaceutical composition, such as a pharmaceutical composition containing an ion pairing agent.
- the 6,8-bis- benzylthio-octanoic acid or a pharmaceutically acceptable salt thereof may be formulated as a pharmaceutical composition for administration to the patient separate from a pharmaceutical composition containing other agent(s) used in the combination therapy, such as docetaxel.
- the methods, compositions, and medical kits for treating prostate cancer provide particular benefits to adult, human male patients suffering from prostate cancer.
- one aspect of the invention provides a method for treating a cancer selected from the group consisting of androgen-dependent prostate cancer and androgen- independent prostate cancer.
- the method comprises administering to a patient in need thereof a therapeutically effective amount of a first therapeutic agent comprising 6,8-bis-benzylthio- octanoic acid or a pharmaceutically acceptable salt thereof, in order to treat the cancer.
- the first therapeutic agent may be administered, for example, by oral or intravenous administration.
- Another aspect of the invention provides a method for treating prostate cancer, where the method comprises administering to a patient in need thereof a therapeutically effective amount of (i) a first therapeutic agent comprising 6,8-bis-benzylthio-octanoic acid or a pharmaceutically acceptable salt thereof and (ii) a second therapeutic agent comprising docetaxel or a pharmaceutically acceptable salt thereof, in order to treat the prostate cancer.
- the method may be characterized according to, for example, the type of prostate cancer, such as an androgen-dependent prostate cancer or an androgen-independent prostate cancer.
- the first therapeutic agent may be administered, for example, by oral or intravenous administration.
- Another aspect of the invention provides a method for treating prostate cancer according to a particular dosing amount.
- the method comprises administering to a patient in need thereof a unit dosage of a first therapeutic agent comprising 6,8-bis-benzylthio-octanoic acid or a pharmaceutically acceptable salt thereof in an amount ranging from about 150 mg/m 2 to about 1500 mg/m 2 , to treat the prostate cancer.
- the method may be characterized according to, for example, the type of prostate cancer, such as an androgen-dependent prostate cancer or an androgen-independent prostate cancer.
- the first therapeutic agent may be administered, for example, by oral or intravenous administration.
- kits for treating a cancer selected from the group consisting of androgen-dependent prostate cancer and androgen-independent prostate cancer comprises (i) a first therapeutic agent comprising 6,8-bis-benzylthio- octanoic acid or a pharmaceutically acceptable salt thereof, and (ii) instructions for treating a cancer selected from the group consisting of androgen-dependent prostate cancer and androgen-independent prostate cancer using said first therapeutic agent.
- the instructions may specify, for example, the route of administration for the first therapeutic agent, such as by oral or intravenous administration.
- kits for treating prostate cancer comprises (i) a first therapeutic agent comprising 6,8-bis-benzylthio-octanoic acid or a pharmaceutically acceptable salt thereof, and (ii) instructions for treating prostate cancer using the first therapeutic agent in combination with a second therapeutic agent comprising docetaxel or a pharmaceutically acceptable salt thereof.
- the instructions may specify, for example, the type of prostate cancer to be treated, such as an androgen-dependent prostate cancer or an androgen-independent prostate cancer.
- the instructions may specify, for example, the route of administration for the first therapeutic agent, such as by oral or intravenous administration.
- kits for treating prostate cancer comprises (i) a first therapeutic agent comprising 6,8-bis-benzylthio-octanoic acid or a pharmaceutically acceptable salt thereof, and (ii) instructions for treating prostate cancer using a dosage of the first therapeutic agent ranging from about 150 mg/m 2 to about 1500 mg/m 2 .
- the instructions may specify, for example, the type of prostate cancer to be treated, such as an androgen-dependent prostate cancer or an androgen-independent prostate cancer.
- the instructions may specify, for example, the route of administration for the first therapeutic agent, such as by oral or intravenous administration.
- Figure 1 is graph a showing the percentage of remaining live PC-3 androgen resistant prostate cancer cells observed following the treatment protocol, as described in Example 2.
- Figure 2 is graph showing observed tumor volume over time after administration of 2.5 mg/kg or 10 mg/kg CPI-613 according to the procedures described in Example 3.
- Figure 3 is graph showing mouse survival time after administration of 2.5 mg/kg or 10 mg/kg CPI-613 according to the procedures described in Example 3.
- Figure 4 depicts the anti-tumor efficacy of oral 6,8-bis-benzylthio-octanoic acid in human non-small cell lung cancer xenografts in mice.
- Figure 5 depicts the anti-tumor efficacy of oral 6,8-bis-benzylthio-octanoic acid in human pancreatic cancer xenografts in mice.
- Figure 6 presents X-ray powder diffraction patterns of solid amorphous dispersion formulations of 6,8-bis-benzylthio-octanoic acid with either Eudragit L100 or hydroxypropyl methylcellulose acetate succinate (HPMCAS-M) (top and middle diffraction patterns, respectively), and crystalline 6,8-bis-benzylthio-octanoic acid (bottom diffraction pattern).
- HPMCAS-M hydroxypropyl methylcellulose acetate succinate
- the invention provides methods, compositions, and medical kits for treating prostate cancer using 6,8-bis-benzylthio-octanoic acid or a pharmaceutically acceptable salt thereof, including combination therapy with docetaxel.
- the prostate cancer may be, for example, characterized as an androgen-dependent prostate cancer or an androgen-independent prostate cancer.
- the 6,8-bis-benzylthio-octanoic acid or a pharmaceutically acceptable salt thereof may be formulated as a pharmaceutical composition, such as a pharmaceutical composition containing an ion pairing agent.
- the 6,8-bis- benzylthio-octanoic acid or a pharmaceutically acceptable salt thereof may be formulated as a pharmaceutical composition for administration to the patient separate from a pharmaceutical composition containing other agent(s) used in the combination therapy, such as docetaxel.
- the methods, compositions, and medical kits for treating prostate cancer provide particular benefits to adult, human male patients suffering from prostate cancer.
- the practice of the present invention employs, unless otherwise indicated, conventional techniques of organic chemistry, pharmacology, and biochemistry. Such techniques are explained in the literature, such as “Comprehensive Organic Synthesis” (B.M. Trost & I. Fleming, eds., 1991-1992); which is incorporated by reference.
- Various aspects of the invention are set forth below in sections; however, aspects of the invention described in one particular section are not to be limited to any particular section. I. DEFINITIONS
- compositions of the present invention may exist in particular geometric or stereoisomeric forms.
- the present invention contemplates all such compounds, including cis- and trans-isomers, R- and L-enantiomers. diastereomers, (D)- isomers, (L)-isomers, the racemic mixtures thereof, and other mixtures thereof, as falling within the scope of the invention.
- the term“patient” refers to organisms to be treated by the methods of the present invention.
- Such organisms preferably include, but are not limited to, mammals (e.g., murines, simians, equines (horses), bovines (cattle), porcines, canines, felines, and the like), and most preferably includes humans.
- the term“treating” includes any effect, e.g., lessening, reducing, modulating, ameliorating or eliminating, that results in the improvement of the condition, disease, disorder, and the like, or ameliorating a symptom thereof.
- treatment can include diminishment of a symptom of a disorder or complete eradication of a disorder.
- treatment can include slowing the progression of a disease, or preventing or delaying its recurrence, such as maintenance treatment to prevent or delay relapse.
- the term“pharmaceutical composition” refers to the combination of an active agent with a carrier, inert or active, making the composition especially suitable for diagnostic or therapeutic use in vivo or ex vivo.
- phrases "pharmaceutically acceptable” is employed herein to refer to those compounds, materials, compositions, and/or dosage forms which are, within the scope of sound medical judgment, suitable for use in contact with the tissues of human beings and animals without excessive toxicity, irritation, allergic response, or other problem or complication, commensurate with a reasonable benefit/risk ratio.
- compositions also can include stabilizers and preservatives.
- carriers, stabilizers, and adjuvants see e.g., Martin,
- salts of the compounds of the present invention may be derived from inorganic or organic acids and bases.
- acids include, but are not limited to, hydrochloric, hydrobromic, sulfuric, nitric, perchloric, fumaric, maleic, phosphoric, glycolic, lactic, salicylic, succinic, toluene-p-sulfonic, tartaric, acetic, citric, methanesulfonic, ethanesulfonic, formic, benzoic, malonic, naphthalene-2-sulfonic, benzenesulfonic acid, and the like.
- Other acids such as oxalic, while not in themselves pharmaceutically acceptable, may be employed in the preparation of salts useful as intermediates in obtaining the compounds of the invention and their pharmaceutically acceptable acid addition salts.
- bases include, but are not limited to, alkali metals (e.g., sodium) hydroxides, alkaline earth metals (e.g. , magnesium), hydroxides, ammonia, and compounds of formula NW3, wherein W is C1-4 alkyl, and the like.
- salts include salts made using the ion pairing agents described in U.S. Patent No. 8,263,653, the entire disclosure of which is incorporated by reference herein. Still further ion pairing agents can be selected with guidance from Handbook of Pharmaceutical Salts Properties, Selection and Use, UIPAC, Wiley -VCH, P.H. Stahl, ed., the entire disclosure of which is incorporated by reference herein.
- salts include acetate, adipate, alginate, aspartate, benzoate, benzenesulfonate, bisulfate, butyrate, citrate, camphorate, camphorsulfonate,
- cyclopentanepropionate digluconate, dodecylsulfate, ethanesulfonate, fumarate, flucoheptanoate, glycerophosphate, hemisulfate, heptanoate, hexanoate, hydrochloride, hydrobromide, hydroiodide, 2-hydroxyethanesulfonate, lactate, maleate, methanesulfonate, 2- naphthalenesulfonate, nicotinate, oxalate, palmoate, pectinate, persulfate, phenylpropionate, picrate, pivalate, propionate, succinate, tartrate, thiocyanate, tosylate, undecanoate, and the like.
- salts include anions of the compounds of the present invention compounded with a suitable cation such as Na + , NH4 1 . and NWV (wherein W is a C 14 alkyl group), and the like.
- a suitable cation such as Na + , NH4 1 . and NWV (wherein W is a C 14 alkyl group), and the like.
- alkyl is art-recognized, and includes saturated aliphatic groups, including straight-chain alkyl groups and branched-chain alkyl groups.
- the pharmaceutically acceptable salts are those prepared from the following acids: hydrochloric, hydrobromic, sulfuric, nitric, phosphoric, maleic, acetic, palicylic, p-toluene sulfonic, tartaric, citric, methane sulfonic, formic, malonic, succinic, naphthalene-2-sulfonic, and benzene sulfonic.
- the pharmaceutically acceptable salts are those prepared from the following acids: hydrochloric, hydrobromic, sulfuric, nitric, phosphoric, maleic, acetic, palicylic, p-toluene sulfonic, tartaric, citric, methane sulfonic, formic, malonic, succinic, naphthalene-2-sulfonic, and benzene sulfonic.
- the pharmaceutically acceptable salts are those prepared from the following acids: hydrochloric, hydrobromic, sulfuric, nitric
- salts are alkaline metal or alkaline earth salts, such as sodium, potassium or calcium salts of a carboxylic acid group.
- salts of the compounds of the present invention are contemplated as being pharmaceutically acceptable.
- salts of acids and bases that are non- pharmaceutically acceptable may also find use, for example, in the preparation or purification of a pharmaceutically acceptable compound.
- compositions and kits are described as having, including, or comprising specific components, or where processes and methods are described as having, including, or comprising specific steps, it is contemplated that, additionally, there are compositions and kits of the present invention that consist essentially of, or consist of, the recited components, and that there are processes and methods according to the present invention that consist essentially of, or consist of, the recited processing steps.
- compositions specifying a percentage are by weight unless otherwise specified. Further, if a variable is not accompanied by a definition, then the previous definition of the variable controls.
- the invention provides methods for treating prostate cancer using 6,8-bis- benzylthio-octanoic acid or a pharmaceutically acceptable salt thereof.
- the therapeutic methods embrace mono-therapy using 6,8-bis-benzylthio-octanoic acid or a pharmaceutically acceptable salt thereof, as well as combination therapy with a second anti-cancer agent, such as docetaxel. The methods are described in more detail below.
- One aspect of the invention provides a method for treating prostate cancer.
- the invention provides a method for treating a cancer selected from the group consisting of androgen-dependent prostate cancer and androgen-independent prostate cancer.
- the method comprises administering to a patient in need thereof a therapeutically effective amount of a first therapeutic agent comprising 6,8-bis-benzylthio-octanoic acid or a pharmaceutically acceptable salt thereof, in order to treat the cancer.
- the method may be further characterized according to one or more features described herein below.
- Another aspect of the invention provides a method for treating prostate cancer comprising administering to a patient in need thereof a therapeutically effective amount of (i) a first therapeutic agent comprising 6,8-bis-benzylthio-octanoic acid or a pharmaceutically acceptable salt thereof and (ii) a second therapeutic agent comprising docetaxel or a pharmaceutically acceptable salt thereof, in order to treat the prostate cancer.
- a first therapeutic agent comprising 6,8-bis-benzylthio-octanoic acid or a pharmaceutically acceptable salt thereof
- a second therapeutic agent comprising docetaxel or a pharmaceutically acceptable salt thereof
- Another aspect of the invention provides a method for treating prostate cancer comprising administering to a patient in need thereof a unit dosage of a first therapeutic agent comprising 6,8-bis-benzylthio-octanoic acid or a pharmaceutically acceptable salt thereof in an amount ranging from about 150 mg/m 2 to about 1500 mg/m 2 , to treat the prostate cancer.
- the method may be further characterized according to one or more features described herein below.
- the above first therapeutic method may be further characterized by additional features, such as the type of cancer treated and whether a second therapeutic agent is administered to the patient.
- the therapeutic method can be further characterized according to the type of cancer to be treated.
- the cancer is androgen-dependent prostate cancer.
- the cancer is androgen-independent prostate cancer.
- the cancer may be further characterized according to whether it is metastatic. In certain embodiments, the cancer is metastatic. Administering a Second Therapeutic Agent
- the therapeutic method may be characterized according to whether a second therapeutic agent is administered to the patient.
- a second therapeutic agent having anti-cancer activity is administered to the patient.
- the above second therapeutic method may be further characterized by additional features, such as the type of prostate cancer treated and the identity of the second therapeutic agent.
- the therapeutic method can be further characterized according to the type of prostate cancer to be treated.
- the prostate cancer is androgen- dependent prostate cancer.
- the prostate cancer is androgen- independent prostate cancer.
- the prostate cancer may be further characterized according to whether it is metastatic. In certain embodiments, the prostate cancer is metastatic.
- the therapeutic method may be further characterized according to the identity of the second therapeutic agent.
- the second therapeutic agent is docetaxel.
- the therapeutic method may be characterized according to features of administration of the second therapeutic agent.
- the therapeutic method may be characterized according to the dose of the second therapeutic agent administered to the patient.
- the second therapeutic agent is administered at a dosage ranging from about 50 mg/m 2 to about 100 mg/m 2 .
- the second therapeutic agent is administered at a dosage of about 75 mg/m 2 .
- the therapeutic method may also be characterized according to the frequency of administration of the second therapeutic agent.
- the second therapeutic agent is administered once per any three week period.
- the therapeutic method may be further characterized according to the route of administration of the second therapeutic agent.
- the second therapeutic agent is administered by intravenous administration.
- the therapeutic method may also be characterized according to the scheduled dosing cycle for administration of the docetaxel.
- the docetaxel is administered on the first day of a three week cycle.
- the docetaxel is administered at a dose from about 50 mg/m 2 to about 100 mg/m 2 on the first day of a three week cycle.
- the docetaxel is administered at a dosage of about 75 mg/m 2 on the first day of a three week cycle.
- the docetaxel is administered as a one hour infusion on the first day of a three week cycle.
- the scheduled cycle is repeated at least once.
- the method of the present invention comprises treatment with up to 10 scheduled cycles.
- first and second therapeutic methods may be further characterized by additional features, such as the unit dosage, timing, and route for administration of the first therapeutic agent.
- the therapeutic methods may be characterized according to features of
- the therapeutic methods may be characterized according to the unit dosage of the first therapeutic agent administered to the patient.
- the first therapeutic agent is administered at a unit dosage ranging from about 150 mg/m 2 to about 1500 mg/m 2 .
- the first therapeutic agent is administered at a unit dosage ranging from about 150 mg/m 2 to about 1300 mg/m 2 .
- the first therapeutic agent is administered at a unit dosage ranging from about 500 mg/m 2 to about 1000 mg/m 2 .
- the first therapeutic agent is administered at a unit dosage of about 500 mg/m 2 .
- the therapeutic methods may be further characterized according to the route of administration of the first therapeutic agent.
- the first therapeutic agent is administered orally.
- the first therapeutic agent is administered by intravenous administration.
- the therapeutic methods may also be characterized according to the frequency of administration of the first therapeutic agent.
- the first therapeutic agent is administered to the patient no more frequently than once per day.
- the first therapeutic agent is administered to the patient twice per week.
- the first therapeutic agent is administered to the patient once per week. In certain other embodiments, the first therapeutic agent is administered to the patient once every two weeks. In certain other embodiments, the first therapeutic agent is administered to the patient no more frequently than once every two weeks.
- the therapeutic method may also be characterized according to the scheduled dosing cycle for administration of the first therapeutic agent.
- the first therapeutic agent is administered in a four week cycle in which the first therapeutic agent is administered twice per week during the first three weeks followed by one week off.
- the first therapeutic agent is administered on days 1, 8, and 15 of a four week cycle.
- the first therapeutic agent is administered on days 1 and 3 of a two week cycle.
- the first therapeutic agent is administered at a dose of about 150 mg/m 2 to about 1500 mg/m 2 twice per week during the first three weeks followed by one week off.
- the first therapeutic agent is administered at a dose of about 150 mg/m 2 to about 1500 mg/m 2 on days 1, 8, and 15 of a four week cycle.
- the first therapeutic agent is administered at a dose of about 150 mg/m 2 to about 1500 mg/m 2 on days 1 and 3 of a two week cycle. In certain embodiments, the first therapeutic agent is administered at a dose of about 500 mg/m 2 to about 100 mg/m 2 twice per week during the first three weeks followed by one week off. In certain embodiments, the first therapeutic agent is administered at a dose of about 500 mg/m 2 to about 100 mg/m 2 on days 1, 8, and 15 of a four week cycle. In certain embodiments, the first therapeutic agent is administered at a dose of about 500 mg/m 2 to about 1000 mg/m 2 on days 1 and 3 of a two week cycle.
- the first therapeutic agent is administered at a dose of about 150 mg/m 2 to about 500 mg/m 2 twice per week during the first three weeks followed by one week off. In certain embodiments, the first therapeutic agent is administered at a dose of about 150 mg/m 2 to about 500 mg/m 2 on days 1, 8, and 15 of a four week cycle. In certain embodiments, the first therapeutic agent is administered at a dose of about 150 mg/m 2 to about 500 mg/m 2 on days 1 and 3 of a two week cycle. In certain embodiments, the first therapeutic agent is administered at a dose of about 500 mg/m 2 twice per week during the first three weeks followed by one week off. In certain embodiments, the first therapeutic agent is administered at a dose of about 500 mg/m 2 on days 1, 8, and 15 of a four week cycle. In certain embodiments, the first therapeutic agent is administered at a dose of about 150 mg/m 2 to about 500 mg/m 2 twice per week during the first three weeks followed by one week off. In certain embodiments, the first therapeutic agent is administered at a dose of about 500 mg
- the first therapeutic agent is administered at a dose of about 500 mg/m 2 on days 1 and 3 of a two week cycle. In certain embodiments, each of the above doses of the first therapeutic agent is administered as a two hour IV infusion. In certain embodiments, the scheduled cycle is repeated at least once. In certain embodiments, the method of the present invention comprises treatment with up to 10 scheduled cycles.
- the first therapeutic agent may be orally administered to the patient. The dose and schedule will vary based on, e.g., the characteristics of the patient’s cancer and whether another therapeutic agent will be administered in combination, and can be readily determined by those of ordinary skill in the art in view of the guidance provided herein.
- the dose and schedule is adapted based on the doses and schedules used intravenously with 6,8-bis- benzylthio-octanoic acid or a pharmaceutically acceptable salt thereof, such as those set forth herein.
- the dose is the maximum tolerated dose.
- An advantage of oral dosing of the first therapeutic agent is that it permits substantially increased dosing flexibility as compared to IV.
- 6,8-bis-benzylthio- octanoic acid is formulated as a 50 mg/mL solution in 1 M (150 mg/mL) aqueous
- triethanolamine which is diluted from 50 mg/mL to as low as 4 mg/mL (e.g., 12.5 mg/mL) with sterile 5% dextrose for injection (D5W) prior to administration as an IV infusion over 30- 120 minutes via a central venous catheter.
- D5W sterile 5% dextrose for injection
- Such an infusion is inconvenient for patients and effectively precludes regimens involving frequent and/or prolonged dosing. Since the half-life of the first therapeutic agent after IV dosing is only about 1-2 hours (Pardee, T.S. et al, Clin Cancer Res. 2014, 20, 5255-64), more frequent and/or prolonged dosing could advantageously be used to increase the patient’s exposure to the first therapeutic agent.
- a possible IV schedule for the treatment of prostate cancer involves administering the first therapeutic agent twice per week for the first three weeks of a four week cycle. If administered orally, the practitioner would have more flexibility with respect to the first therapeutic agent dose and schedule.
- the first therapeutic agent could be orally administered in a single daily dose twice per week for the first three weeks of a four week cycle as in the IV schedule.
- the first therapeutic agent could be administered in two or more (e.g., three, four, or five) divided doses on two days per week for the first three weeks of a four week cycle and/or the first therapeutic agent could be administered on fewer or additional days of the cycle, up to and including every day.
- Another advantage of oral dosing is that it makes maintenance therapy feasible.
- a patient who is treated successfully with first line therapy - with or without the first therapeutic agent - and whose cancer is in partial or complete remission may be treated orally with the first therapeutic agent on a chronic basis in order to delay or prevent recurrence.
- the maintenance treatment may involve, for example, one, two, three, four, or five doses per day of the first therapeutic agent on a regular basis, such as daily or weekly.
- the first therapeutic agent is orally administered at a dose of about 1 mg to about 10,000 mg on each day it is administered.
- the daily dose may be administered in one dose or divided into two or more doses, such as three, four, or five doses.
- the daily dose is about 10 mg to about 7,500 mg.
- the daily dose is about 100 mg to about 5,000 mg.
- the daily dose is about 200 mg to about 4,000 mg.
- the daily dose is about 300 mg to about 3,000 mg.
- the daily dose is about 400 mg to about 2,500 mg.
- the daily dose is about 500 mg to about 2,000 mg.
- the daily dose is about 100 mg, 200 mg, 300 mg, 400 mg, 500 mg, 600 mg, 700 mg, 800 mg, 900 mg, 1,000 mg, 1,250 mg, 1,500 mg, 1,750 mg, 2,000 mg, 2,500 mg, 3,000 mg, 3,500 mg, 4,000 mg, 4,500 mg, 5,000 mg, 6,000 mg, 7,000 mg, 8,000 mg, 9,000 mg, or 10,000 mg.
- a dosing cycle is repeated at least once.
- the method of the present invention comprises treatment with two cycles or more.
- the method of the present invention comprises treatment with three cycles or more.
- the method of the present invention comprises treatment with four cycles or more.
- the method of the present invention comprises treatment with five cycles or more.
- the method of the present invention comprises treatment with six cycles or more.
- the method of the present invention comprises treatment with seven cycles or more.
- the method of the present invention comprises treatment with eight cycles or more.
- the method of the present invention comprises treatment with nine cycles or more.
- the method of the present invention comprises treatment with ten cycles or more. In certain embodiments, the method of the present invention comprises regular treatment with the first therapeutic agent, including on a daily or weekly basis, for an extended period of time, such as one year, two years, three years, or longer.
- the above third therapeutic method may be further characterized by additional features, such as the amount, timing, and route for administration of the unit dosage.
- the therapeutic method may be characterized according to features of administration of the unit dosage. For instance, in certain embodiments, the therapeutic methods may be characterized according to the amount of the unit dosage administered to the patient.
- the unit dosage is from about 500 mg/m 2 to about 1000 mg/m 2 of 6,8-bis-benzylthio-octanoic acid or a pharmaceutically acceptable salt thereof. In certain other embodiments, the unit dosage is about 500 mg/m 2 of 6,8-bis-benzylthio-octanoic acid or a pharmaceutically acceptable salt thereof.
- the therapeutic method may also be characterized according to the frequency of administration of the unit dosage. For example, in certain embodiments, the unit dosage is administered to the patient no more frequently than once per day. In certain embodiments, the unit dosage is administered to the patient twice per week. In certain other embodiments, the unit dosage is administered to the patient once per week. In certain other embodiments, the unit dosage is administered to the patient once every two weeks. In certain other embodiments, the unit dosage is administered to the patient no more frequently than once every two weeks.
- the therapeutic method may be further characterized according to the route of administration of the unit dosage.
- the unit dosage is administered intravenously to the patient.
- the unit dosage is administered orally to the patient.
- the therapeutic method may also be characterized according to the scheduled dosing cycle for administration of the 6,8-bis-benzylthio-octanoic acid or pharmaceutically acceptable salt thereof.
- the 6,8-bis-benzylthio-octanoic acid or pharmaceutically acceptable salt thereof is administered in a four week cycle in which the 6,8- bis-benzylthio-octanoic acid or pharmaceutically acceptable salt thereof is administered twice per week during the first three weeks followed by one week off.
- the 6,8-bis-benzylthio-octanoic acid or pharmaceutically acceptable salt thereof is administered on days 1, 8, and 15 of a four week cycle.
- the 6,8-bis-benzylthio-octanoic acid or pharmaceutically acceptable salt thereof is administered on days 1 and 3 of a two week cycle. In certain embodiments, the 6,8-bis-benzylthio-octanoic acid or pharmaceutically acceptable salt thereof is administered at a dose of about 150 mg/m 2 to about 1500 mg/m 2 twice per week during the first three weeks followed by one week off. In certain embodiments, the 6,8-bis-benzylthio-octanoic acid or pharmaceutically acceptable salt thereof is administered at a dose of about 150 mg/m 2 to about 1500 mg/m 2 on days 1, 8, and 15 of a four week cycle.
- the 6,8-bis-benzylthio-octanoic acid or pharmaceutically acceptable salt thereof is administered at a dose of about 150 mg/m 2 to about 1500 mg/m 2 on days 1 and 3 of a two week cycle. In certain embodiments, the 6,8-bis-benzylthio-octanoic acid or
- the 6,8-bis-benzylthio-octanoic acid or pharmaceutically acceptable salt thereof is administered at a dose of about 500 mg/m 2 to about 100 mg/m 2 twice per week during the first three weeks followed by one week off.
- the 6,8-bis-benzylthio-octanoic acid or pharmaceutically acceptable salt thereof is administered at a dose of about 500 mg/m 2 to about 100 mg/m 2 on days 1, 8, and 15 of a four week cycle.
- the 6,8-bis-benzylthio-octanoic acid or pharmaceutically acceptable salt thereof is administered at a dose of about 500 mg/m 2 to about 1000 mg/m 2 on days 1 and 3 of a two week cycle.
- the 6,8-bis-benzylthio-octanoic acid or pharmaceutically acceptable salt thereof is administered at a dose of about 150 mg/m 2 to about 500 mg/m 2 twice per week during the first three weeks followed by one week off. In certain embodiments, the 6,8-bis-benzylthio-octanoic acid or pharmaceutically acceptable salt thereof is administered at a dose of about 150 mg/m 2 to about 500 mg/m 2 on days 1, 8, and 15 of a four week cycle. In certain embodiments, the 6,8-bis-benzylthio-octanoic acid or pharmaceutically acceptable salt thereof is administered at a dose of about 150 mg/m 2 to about 500 mg/m 2 on days 1 and 3 of a two week cycle.
- the 6,8-bis- benzylthio-octanoic acid or pharmaceutically acceptable salt thereof is administered at a dose of about 500 mg/m 2 twice per week during the first three weeks followed by one week off. In certain embodiments, the 6,8-bis-benzylthio-octanoic acid or pharmaceutically acceptable salt thereof is administered at a dose of about 500 mg/m 2 on days 1, 8, and 15 of a four week cycle. In certain embodiments, the 6,8-bis-benzylthio-octanoic acid or pharmaceutically acceptable salt thereof is administered at a dose of about 500 mg/m 2 on days 1 and 3 of a two week cycle.
- each of the above doses of the 6,8-bis-benzylthio-octanoic acid or pharmaceutically acceptable salt thereof is administered as a two hour IV infusion.
- the scheduled cycle is repeated at least once.
- the method of the present invention comprises treatment with up to 10 scheduled cycles.
- first, second, and third therapeutic methods may be further characterized by additional features, such as the patients to be treated and the form of the first therapeutic agent.
- the therapeutic methods may be further characterized according to the patient to be treated.
- the patient is a human being.
- the patient is an adult human.
- the patient is at least partially refractory to docetaxel.
- the methods may be further characterized according to the form of the first therapeutic agent.
- the first therapeutic agent is 6,8-bis- benzylthio-octanoic acid or a pharmaceutically acceptable salt thereof.
- the first therapeutic agent is 6,8-bis-benzylthio-octanoic acid.
- the first therapeutic agent may be formulated in a pharmaceutical composition.
- the first therapeutic agent is administered in the form of a pharmaceutical composition comprising 6,8-bis-benzylthio-octanoic acid and a pharmaceutically acceptable carrier.
- the first therapeutic agent is administered in the form of a pharmaceutical composition comprising 6,8-bis-benzylthio-octanoic acid and an ion pairing agent.
- the first therapeutic agent is administered in the form of a pharmaceutical composition comprising 6,8-bis-benzylthio-octanoic acid and triethanolamine.
- the pharmaceutical composition further comprises dextrose and water.
- Exemplary ion pairing agents that may be used include, for example, a tertiary amine (such as triethanolamine), other amines such as diethanolamine, monoethanolamine, mefenamic acid and tromethamine, and combinations thereof.
- the ion pairing agent is an organic Bronsted base.
- the ion pairing agent is an amine compound.
- the ion pairing agent is a monoalkylamine, dialkylamine, trialkylamine, amino-substituted aliphatic alcohol, hydroxymonoalkylamine, hydroxy dialkylamine, hydroxytrialkylamine, amino-substituted heteroaliphatic alcohol, alkyldiamine, substituted alkyldiamine, or optionally substituted heteroaryl group containing at least one ring nitrogen atom.
- Additional exemplary ion pairing agents include, for example, polyethyleneimine, polyglutamic acid, ammonia, L-arginine, benethamine benzathine, betaine, calcium hydroxide, choline, deanol, diethanolamine(2,2’-iminobis(ethanol)), diethylamine, 2-(diethylamino)- ethanol, ethanolamine, ethylenediamine, N-methyl-glucamine, hydrabamine, lH-imidazole, lysine, magnesium hydroxide, 4-(2-hydroxyethyl)-morpholine, piperazine, potassium hydroxide, 1 -(2-hydroxy ethyl)-pyrrolidine, sodium hydroxide, triethanolamine (2, 2’, 2”- nitrilotris(ethanol)), tromethamine, and zinc hydroxide.
- the ion pairing agent is diisopropanolamine, 3-amino- 1 -propanol, meglumine, morpholine, pyridine, niacinamide, tris(hydroxymethyl)aminomethane, 2-((2-dimethylamino)ethoxy)ethanol, 2- (dimethylamino)ethanol, l-(2-hydroxyethyl)pyrrolidine, or ammonium hydroxide.
- the ion pairing agent is an alkali metal hydroxide or alkaline earth metal hydroxide, such as, for example, cesium hydroxide.
- the first therapeutic agent has a purity of at least about 50% (w/w). In certain embodiments, the first therapeutic agent has a purity of at least about 60% (w/w). In certain embodiments, the first therapeutic agent has a purity of at least about 70% (w/w). In certain embodiments, the first therapeutic agent has a purity of at least about 80% (w/w). In certain embodiments, the first therapeutic agent has a purity of at least about 90% (w/w). In certain embodiments, the first therapeutic agent has a purity of at least about 95% (w/w). In certain embodiments, the first therapeutic agent has a purity of at least about 96%
- the first therapeutic agent has a purity of at least about 97%
- the first therapeutic agent has a purity of at least about 98%
- the first therapeutic agent has a purity of at least about 99%
- the first therapeutic agent is administered in the form of a pharmaceutical composition comprising 6,8-bis-benzylthio-octanoic acid and
- Such pharmaceutical compositions may be further characterized according to the mole ratio of the triethanolamine to 6,8-bis-benzylthio-octanoic acid.
- the mole ratio of triethanolamine to 6,8-bis-benzylthio-octanoic acid is in the range of about 10: 1 to about 1: 10, about 10: 1 to about 5: 1, or about 8: 1.
- the mole ratio of triethanolamine to 6,8-bis-benzylthio-octanoic acid is about 8: 1.
- a therapeutic agent e.g., 6,8-bis-benzylthio-octanoic acid or a pharmaceutically acceptable salt thereof, is delivered to the patient in a therapeutically effective amount.
- the therapeutically effective amount of a therapeutic agent may vary with the activity of the specific agent employed; the metabolic stability and length of action of that agent; the species, age, body weight, general health, dietary status, sex and diet of the subject; the mode and time of administration; rate of excretion; drug combination, if any; and extent of presentation and/or severity of the particular condition being treated.
- the precise dosage can be determined, may involve one or several administrations per day, in whichever order is necessary or desirable, to yield the desired results, and the dosage may be adjusted by the individual practitioner to achieve a desired effect.
- the treatment may involve one or several administrations on one or more days, and the dosage may be adjusted by the individual practitioner to achieve a desired effect.
- the dosage amount of the agent(s) used should be sufficient to interact solely with disease cells (e.g., tumor cells), leaving normal cells comparatively unharmed (e.g., essentially unharmed).
- the dosage amount may be administered in a single dose or in the form of individual divided doses, such as from one to four or more times per day.
- the daily dosage amount is administered in a single dose.
- Components in a combination therapy may be administered in a particular order and/or according to a treatment cycle, such as on the same or different days. For example, in certain embodiments, at least one dose of the first therapeutic agent is administered to the patient prior to administering the second therapeutic agent, such as on an earlier day in a treatment cycle.
- active components of the combination therapy may be administered on the same day of a treatment cycle, for example being co-administered simultaneously.
- at least one dose of a second therapeutic agent is administered to the patient prior to administering the 6,8-bis-benzylthio-octanoic acid or a pharmaceutically acceptable salt thereof, such as on an earlier day in a treatment cycle.
- active components of the combination therapy may be co-administered in a predetermined manner, ratio, and order of addition so as to comprise a treatment cycle.
- treatment cycles may be repeated in order to maximize benefit to the patient.
- the therapeutic method of the present invention may be further characterized by the efficacy and safety of the treatment.
- the method provides an acceptable safety profile, with the benefit of treatment outweighing the risk.
- the method of the present invention preferably provides an overall response rate of at least about 10%, a duration of response of at least about 1 month, progression-free survival (PFS) of at least about 1 month, and/or overall survival (OS) of at least about 1 month.
- the phase II or phase III clinical trial comprises at least 15 patients. More preferably, the phase II or phase III clinical trial comprises at least 20 patients. More preferably, the phase II or phase III clinical trial comprises at least 25 patients.
- the phase II or phase III clinical trial comprises at least 50 patients. More preferably, the phase II or phase III clinical trial comprises at least 100 patients. More preferably, the phase II or phase III clinical trial comprises at least 200 patients. More preferably, the phase II or phase III clinical trial comprises at least 300 patients. More preferably, the phase II or phase III clinical trial comprises at least 400 patients. More preferably, the phase II or phase III clinical trial comprises at least 500 patients.
- the method of the present invention provides an overall response rate of at least about 20% in patients. More preferably, the method of the present invention provides an overall response rate of at least about 30%. More preferably, the method of the present invention provides an overall response rate of at least about 40%.
- the method of the present invention provides an overall response rate of at least about 50%. More preferably, the method of the present invention provides an overall response rate of at least about 60%. More preferably, the method of the present invention provides an overall response rate of at least about 70%. More preferably, the method of the present invention provides an overall response rate of at least about 80%. More preferably, the method of the present invention provides an overall response rate of at least about 90%.
- the method of the present invention provides a duration of response, PFS, and/or OS of at least about 2 months. Preferably, the method of the present invention provides a duration of response, PFS, and/or OS of at least about 3 months.
- the method of the present invention provides a duration of response, PFS, and/or OS of at least about 4 months.
- the method of the present invention provides a duration of response, PFS, and/or OS of at least about 5 months.
- the method of the present invention provides a duration of response, PFS, and/or OS of at least about 6 months.
- the method of the present invention provides a duration of response, PFS, and/or OS of at least about 7 months.
- the method of the present invention provides a duration of response, PFS, and/or OS of at least about 8 months.
- the method of the present invention provides a duration of response, PFS, and/or OS of at least about 9 months.
- the method of the present invention provides a duration of response, PFS, and/or OS of at least about 10 months.
- the method of the present invention provides a duration of response, PFS, and/or OS of at least about 11 months.
- the method of the present invention provides a duration of response, PFS, and/or OS of at least about 12 months.
- the method of the present invention provides a duration of response, PFS, and/or OS of at least about 14 months.
- the method of the present invention provides a duration of response, PFS, and/or OS of at least about 16 months.
- the method of the present invention provides a duration of response, PFS, and/or OS of at least about 18 months.
- the method of the present invention provides a duration of response, PFS, and/or OS of at least about 20 months.
- the method of the present invention provides a duration of response, PFS, and/or OS of at least about 24 months.
- the overall response rate, duration of response, and progression-free survival mentioned above are measured in a phase II clinical trial.
- the overall response rate, duration of response, and progression-free survival mentioned above are measured in a phase III clinical trial.
- the methods desirably administer a therapeutically effective amount of the indicated compound(s) to the patient.
- a therapeutically effective amount can be determined based on guidance herein and may, for instance, be an amount of a compound sufficient to inhibit, halt, or cause an improvement in a disorder or condition being treated in a particular subject or subject population.
- a therapeutically effective amount can be an amount of drug sufficient to slow the progression of a disease, or to prevent or delay its recurrence, such as maintenance treatment to prevent or delay relapse.
- a therapeutically effective amount can be determined experimentally in a laboratory or clinical setting, or may be the amount required by the guidelines of the United States Food and Drug Administration, or equivalent foreign agency, for the particular disease and subject being treated. It should be appreciated that determination of proper dosage forms, dosage amounts, and routes of administration is within the level of ordinary skill in the pharmaceutical and medical arts.
- kits containing a therapeutic agent and/or pharmaceutical composition described herein, along with instructions for using the kits to treat a disorder described herein.
- the medical kit comprises (i) a first therapeutic agent comprising 6,8-bis-benzylthio-octanoic acid or a pharmaceutically acceptable salt thereof, and (ii) instructions for treating a cancer selected from the group consisting of androgen-dependent prostate cancer and androgen-independent prostate cancer using said first therapeutic agent.
- the medical kit may be further characterized according to one or more of the features described herein in connection with the First Therapeutic Method.
- kits for treating prostate cancer comprising (i) a first therapeutic agent comprising 6,8-bis-benzylthio-octanoic acid or a pharmaceutically acceptable salt thereof, and (ii) instructions for treating prostate cancer using the first therapeutic agent in combination with a second therapeutic agent comprising docetaxel or a pharmaceutically acceptable salt thereof.
- the medical kit may be further characterized according to one or more of the features described herein in connection with the Second Therapeutic Method.
- kits for treating prostate cancer comprising (i) a first therapeutic agent comprising 6,8-bis-benzylthio-octanoic acid or a pharmaceutically acceptable salt thereof, and (ii) instructions for treating prostate cancer using a dosage of the first therapeutic agent ranging from about 500 mg/m 2 to about 1000 mg/m 2 .
- the medical kit may be further characterized according to one or more of the features described herein in connection with the Third Therapeutic Method.
- Therapeutic agents described herein may be formulated as a pharmaceutical composition comprising one or more therapeutic agents and a pharmaceutically acceptable carrier.
- the first therapeutic agent can be formulated as a pharmaceutical composition that, for example, optionally further contains a further anti-cancer agent.
- a pharmaceutical composition that contains both a first therapeutic agent and a second therapeutic agent may be referred to as a co-formulated composition.
- a therapeutic agent may be formulated as a pharmaceutically-acceptable oil; liposome; oil-water or lipid-oil-water emulsion or nanoemulsion; liquid; or salt, crystalline form, or other solid form delivered in a tablet or capsule.
- the therapeutic agent may be combined with a pharmaceutically-acceptable carrier or excipient therefor.
- pharmaceutically- acceptable carriers are well known in the art and include those conventionally used in pharmaceutical compositions, such as salts, lipids, buffers, chelating agents, flavorants, colorants, preservatives, absorption promoters to enhance bioavailability, antimicrobial agents, and combinations thereof, optionally in combination with other therapeutic ingredients.
- compositions may be specially formulated for administration in solid or liquid form, including those adapted for parenteral administration, for example, by subcutaneous, intramuscular, intravenous or epidural injection as, for example, a sterile solution or suspension, or sustained-release formulation.
- Methods of preparing pharmaceutical formulations or pharmaceutical compositions include the step of bringing into association a compound of the present invention with the carrier and, optionally, one or more accessory ingredients.
- the formulations are prepared by uniformly and intimately bringing into association a compound of the present invention with liquid carriers, or finely divided solid carriers, or both, and then, if necessary, shaping the product.
- compositions of this invention suitable for parenteral administration comprise one or more compounds of the invention in combination with one or more pharmaceutically-acceptable sterile isotonic aqueous or nonaqueous solutions, dispersions, suspensions or emulsions, or sterile powders which may be reconstituted into sterile injectable solutions or dispersions just prior to use, which may contain sugars, alcohols, antioxidants, buffers, bacteriostats, solutes which render the formulation isotonic with the blood of the intended recipient or suspending or thickening agents.
- one or more of the therapeutic agents are administered by intraparenteral administration.
- one or more of the therapeutic agents are formulated for inhalational, oral, topical, transdermal, nasal, ocular, pulmonary, rectal, transmucosal, intravenous, intramuscular, subcutaneous, intraperitoneal, intrathoracic, intrapleural, intrauterine, intratumoral, or infusion methodologies or administration, or combinations of any thereof, in the form of aerosols, sprays, powders, gels, lotions, creams, suppositories, ointments, and the like.
- other additives known in the art may be included to impart the desired consistency and other properties to the formulation.
- the pharmaceutical composition comprising the first therapeutic agent is an oral dosage form, such as a dry oral dosage form.
- the pharmaceutical composition comprising the first therapeutic agent is an oral dosage form chosen from tablet, pill, capsule, caplet, powder, granule, solution, suspension, and gel.
- Oral dosage forms may include pharmaceutically acceptable excipients, such as carriers, diluents, stabilizers, plasticizers, binders, glidants, disintegrants, bulking agents, lubricants, plasticizers, colorants, film formers, flavoring agents, preservatives, dosing vehicles, and any combination of any of the foregoing.
- Pharmaceutically acceptable excipients are determined in part by the particular composition being administered, as well as by the particular dosing schedule.
- compositions of the present invention there is a wide variety of suitable formulations of pharmaceutical compositions of the present invention (see, e.g., Remington: The Science and Practice of Pharmacy, 20th ed., Gennaro et al. Eds., Lippincott Williams and Wilkins, 2000).
- the oral pharmaceutical composition comprising the first therapeutic agent will generally include at least one inert excipient.
- Excipients include pharmaceutically compatible binding agents, lubricants, wetting agents, disintegrants, and the like. Tablets, pills, capsules, troches and the like can contain any of the following excipients, or compounds of a similar nature: a binder such as microcrystalline cellulose, gum tragacanth or gelatin; an excipient such as starch or lactose, a dispersing agent such as alginic acid, Primogel, or com starch; a lubricant such as magnesium stearate; a glidant such as colloidal silicon dioxide; a sweetening agent such as sucrose or saccharin; or a flavoring agent such as peppermint, methyl salicylate, or orange flavoring.
- a binder such as microcrystalline cellulose, gum tragacanth or gelatin
- an excipient such as starch or lactose, a dispersing agent such
- the dosage unit form When the dosage unit form is a capsule, it can contain a liquid excipient such as a fatty oil.
- dosage unit forms can contain various other materials that modify the physical form of the dosage unit, for example, coatings of sugar, shellac, or enteric agents.
- a syrup may contain, in addition to the active compounds, sucrose as a sweetening agent and certain preservatives, dyes, colorings, and flavorings.
- the oral pharmaceutical composition comprising the first therapeutic agent comprises an excipient in an amount of about 5% to about 99%, such as about 10% to about 85%, by weight of the composition, with the first therapeutic agent comprising the remainder.
- pharmaceutically acceptable excipients comprise about 20% to about 80% of the total weight of the composition.
- the pharmaceutical composition comprises the first therapeutic agent in an amount of at least about 40% by weight of the composition, with one or more excipients comprising the remainder. In certain embodiments, the pharmaceutical composition comprises the first therapeutic agent in an amount of at least about 50% by weight of the composition. In certain embodiments, the pharmaceutical composition comprises the first therapeutic agent in an amount of at least about 60% by weight of the composition. In certain embodiments, the pharmaceutical composition comprises the first therapeutic agent in an amount of at least about 70% by weight of the composition. In certain embodiments, the pharmaceutical composition comprises the first therapeutic agent in an amount of at least about 80% by weight of the composition. In certain embodiments, the pharmaceutical composition comprises the first therapeutic agent in an amount of at least about 90% by weight of the composition.
- Diluents for solid oral pharmaceutical compositions comprising the first therapeutic agent include, but are not limited to, microcrystalline cellulose (e.g. AVICEL®), microfine cellulose, lactose, starch, pregelatinized starch, calcium carbonate, calcium sulfate, sugar, dextrates, dextrin, dextrose, dibasic calcium phosphate dihydrate, tribasic calcium phosphate, kaolin, magnesium carbonate, magnesium oxide, maltodextrin, mannitol, polymethacrylates (e.g. Eudragit), potassium chloride, powdered cellulose, sodium chloride, sorbitol and talc.
- microcrystalline cellulose e.g. AVICEL®
- microfine cellulose e.g. AVICEL®
- lactose e.g. AVICEL®
- starch pregelatinized starch
- calcium carbonate calcium sulfate
- sugar dextrates
- dextrin dextrin
- Binders for solid oral pharmaceutical compositions comprising the first therapeutic agent include, but are not limited to, acacia, tragacanth, sucrose, glucose, alginic acid, carbomer (e.g. Carbopol), carboxymethylcellulose sodium, dextrin, ethyl cellulose, gelatin, guar gum, hydrogenated vegetable oil, hydroxyethyl cellulose, hydroxypropyl cellulose (e.g. KLUCEL®), hydroxypropyl methyl cellulose (e.g. METHOCEL®), liquid glucose, magnesium aluminum silicate, maltodextrin, methylcellulose, polymethacrylates, povidone (e.g.
- the pharmaceutical composition comprises a binder in an amount of about 0.5% to about 25%, such as about 0.75% to about 15%, by weight of the composition. In certain embodiments, the pharmaceutical composition comprises a binder in an amount of about 1% to about 10% by weight of the composition.
- the dissolution rate of a compacted solid pharmaceutical composition in a patient's stomach may be increased by the addition of a disintegrant to the composition comprising the first therapeutic agent.
- Disintegrants include, but are not limited to, alginic acid, carboxymethylcellulose calcium, carboxymethylcellulose sodium (e.g. AC -DI-SOL®, PRIMELLOSE®), colloidal silicon dioxide, croscarmellose sodium, crospovidone (e.g.
- the pharmaceutical composition comprises a disintegrant in an amount of about 0.2% to about 30%, such as about 0.2% to about 10%, by weight of the composition. In certain embodiments, the pharmaceutical composition comprises a disintegrant in an amount of about 0.2% to about 5% by weight of the composition.
- the oral pharmaceutical composition comprising the first therapeutic agent optionally comprises one or more pharmaceutically acceptable wetting agents.
- Such wetting agents are preferably selected to maintain the API in close association with water, a condition that is believed to improve bioavailability of the composition.
- surfactants that can be used as wetting agents include quaternary ammonium compounds, for example benzalkonium chloride, benzethonium chloride and cetylpyridinium chloride, dioctyl sodium sulfosuccinate, polyoxyethylene alkylphenyl ethers, for example nonoxynol 9, nonoxynol 10, and octoxynol 9, poloxamers (polyoxyethylene and polyoxypropylene block copolymers), polyoxyethylene fatty acid glycerides and oils, for example polyoxyethylene, caprylic/capric mono- and diglycerides (e.g., LabrasolTM of Gattefosse), polyoxyethylene castor oil and polyoxyethylene hydrogenated castor oil;
- polyoxyethylene stearate polyoxyethylene sorbitan esters
- polysorbate 20 and polysorbate 80 e.g., TweenTM 80 of ICI
- propylene glycol fatty acid esters for example propylene glycol laurate (e.g., LauroglycolTM of Gattefosse)
- sodium lauryl sulfate fatty acids and salts thereof, for example oleic acid, sodium oleate and triethanolamine oleate
- glyceryl fatty acid esters for example glyceryl monostearate
- sorbitan esters for example sorbitan monolaurate, sorbitan monooleate, sorbitan monopalmitate and sorbitan monostearate, tyloxapol, and mixtures thereof.
- the pharmaceutical composition comprising the first therapeutic agent comprises a wetting agent in an amount of about 0.25% to about 15%, such as about 0.4% to about 10%, by weight of the composition. In certain embodiments, the pharmaceutical composition comprises a wetting agent in an amount of about 0.5% to about 5% by weight of the composition. In certain embodiments, the pharmaceutical composition comprises a wetting agent that is an anionic surfactant. In certain embodiments, the pharmaceutical composition comprises sodium lauryl sulfate as a wetting agent. In certain embodiments, the pharmaceutical composition comprises sodium lauryl sulfate in an amount of about 0.25% to about 7%, such as about 0.4% to about 4%, by weight of the composition. In certain embodiments, the pharmaceutical composition comprises sodium lauryl sulfate in an amount of about 0.5% to about 2% by weight of the composition.
- Lubricants e.g., anti-adherents or glidants
- Excipients that may function as lubricants include, but are not limited to, colloidal silicon dioxide, magnesium trisilicate, powdered cellulose, starch, talc and tribasic calcium phosphate.
- Suitable lubricants further include glyceryl behapate (e.g., CompritolTM 888 of Gattefosse); stearic acid and salts thereof, including magnesium, calcium and sodium stearates; zinc stearate; glyceryl monostearate; glyceryl palmitostearate; hydrogenated castor oil;
- glyceryl behapate e.g., CompritolTM 888 of Gattefosse
- an oral pharmaceutical composition comprising the first therapeutic agent comprises a lubricant in an amount of about 0.1% to about 10%, such as about 0.2% to about 8%, by weight of the composition.
- the pharmaceutical composition comprises a lubricant in an amount of about 0.25% to about 5% by weight of the composition.
- the pharmaceutical composition comprises magnesium stearate as a lubricant.
- the pharmaceutical composition comprises colloidal silicon dioxide.
- the pharmaceutical composition comprises talc.
- the composition comprises magnesium stearate or talc in an amount of about 0.5% to about 2% by weight of the composition.
- Flavoring agents and flavor enhancers make the oral dosage form comprising the first therapeutic agent more palatable to the patient.
- Common flavoring agents and flavor enhancers for pharmaceutical products include maltol, vanillin, ethyl vanillin, menthol, citric acid, fumaric acid ethyl maltol, and tartaric acid.
- compositions may also be colored using any pharmaceutically acceptable colorant to improve their appearance and/or facilitate patient identification of the product and unit dosage level.
- the solid oral compositions comprising the first therapeutic agent of the present invention include powders, granulates, aggregates and compacted compositions.
- the dosages may be conveniently presented in unit dosage form and prepared by any of the methods well-known in the pharmaceutical arts. Dosage forms include solid dosage forms like tablets, pills, powders, caplets, granules, capsules, sachets, troches and lozenges.
- the pharmaceutical composition comprising the first therapeutic agent is a tablet.
- the pharmaceutical composition comprising the first therapeutic agent is a spray-dried dispersion.
- the pharmaceutical composition comprising the first therapeutic agent is a spray-dried dispersion comprising at least one polymer chosen from polyacrylate, polymethacrylate, poly(vinylpyrrolidone), hydroxypropyl methyl cellulose (HPMC), cellulose acetate phthalate (CAP), and hydroxypropyl
- the pharmaceutical composition comprising the first therapeutic agent is a spray-dried dispersion comprising at least one polymer chosen from Eudragit L100, poly(vinylpyrrolidone), hydroxypropyl methyl cellulose (HPMC), cellulose acetate phthalate (CAP), and hydroxypropyl methylcellulose acetate succinate (HPMCAS-M).
- the pharmaceutical composition comprising the first therapeutic agent is a spray-dried dispersion comprising at least one polymer chosen from Eudragit L100, poly(vinylpyrrolidone) viscosity grade K30 (PVP K30), hydroxypropyl methyl cellulose (HPMC), cellulose acetate phthalate (CAP), and
- the pharmaceutical composition comprising the first therapeutic agent is a spray-dried dispersion comprising at least one polymer chosen from Eudragit L100 and hydroxypropyl
- the pharmaceutical composition comprising the first therapeutic agent is a spray-dried dispersion comprising Eudragit L100. In certain embodiments, the pharmaceutical composition comprising the first therapeutic agent is a spray-dried dispersion comprising hydroxypropyl methylcellulose acetate succinate (HPMCAS-M).
- formulations of the invention may be buffered by the addition of suitable buffering agents.
- the oral pharmaceutical composition comprising the first therapeutic agent of the present invention is a unit dose composition.
- the pharmaceutical composition contains about 1 mg to about 5000 mg of the first therapeutic agent.
- the pharmaceutical composition contains about 100 mg to about 3000 mg of the first therapeutic agent.
- the pharmaceutical composition contains about 200 mg to about 2000 mg of the first therapeutic agent.
- the pharmaceutical composition contains about 50 mg, 100 mg, 200 mg, 300 mg, 400 mg, 500 mg, 600 mg, 700 mg, 800 mg, 900 mg, 1000 mg, 1100 mg, 1200 mg, 1300 mg, 1400 mg, 1500 mg, 1600 mg, 1700 mg, 1800 mg, 1900 mg, 2000 mg, 2500 mg, or 3000 mg of first therapeutic agent.
- the pharmaceutical composition contains about 300 mg, 500 mg, 700 mg, or 1000 mg of the first therapeutic agent.
- the pharmaceutical composition of the present invention comprises an emulsion, particle, or gel as described in U.S. Patent No. 7,220,428.
- the pharmaceutical composition is a solid or liquid formulation having from about 0.1% to about 75% w/w lipids or fatty acid components.
- the formulation contains about 0.1% to about 15% w/v lipids and fatty acid components.
- the fatty acid component comprises saturated or unsaturated C4, C5, C6, C7, C8, C9, C10, Cl 1, or C12 fatty acids and/or salts of such fatty acids.
- Lipids may include cholesterol and analogs thereof.
- a panel of five prostate cancer cell lines was treated with 6,8-te-benzylthio- octanoic acid in vitro.
- prostate cancer cells were treated with 6,8-te-benzylthio- octanoic acid while in RPMI supplemented with 10% FBS for 60 hrs.
- Cell survival was assayed with CellTiter 96® Aqueous One Solution Cell Proliferation Assay (Promega). EC 50 values were calculated. Part II - Results
- CPI-613 was highly effective at killing prostate cancer cells in vitro.
- EC 50 values reflecting the ability of 6,8-to-benzylthio-octanoic acid to kill the indicated prostate cancer cell lines are provided in Table 1 below.
- PC-3 androgen resistant prostate cancer cells were treated with subtherapeutic doses of (i) 6.8-/i/.v-benzylthio-octanoic acid, (ii) docetaxel, or (iii) 6,8-te-benzylthio-octanoic acid and docetaxel.
- PC-3 androgen resistant prostate cancer cells were treated for 44 hours in RPMI+l0% FBS with (i) a 50 mM solution of 6,8-to-benzylthio-octanoic acid alone, (ii) either a 2 nM solution of docetaxel or a 200 nM solution of docetaxel alone, or (iii) a combination of 6,8-te-benzylthio-octanoic acid and docetaxel at the foregoing stated amounts.
- Prostate cancer cell survival was determined using CellTiterGLO (Promega).
- the CPI-613 was formulated in the same manner as it was for clinical trials. Tumors were measured using a caliper. Tumor volume was recorded over four weeks of therapy.
- mice survival of the mice was measured as weeks on therapy in groups of mice treated with 0 mg/kg, 2.5 mg/kg, or 10 mg/kg of 6,8 -bis- benzylthio-octanoic acid; experimental endpoints included death, tumor ulceration, or tumor volume reaching 750 mm 3 .
- a graph of tumor volume over time is provided in Figure 2. Tumor growth was significantly slower in groups of mice treated with CPI-613. A graph showing mouse survival time is provided in Figure 3 (same mice as in Figure 2). Both groups of mice treated with 6,8- /i/.v-benzylthio-octanoic acid demonstrated a significant decrease in tumor growth and increased survival without toxicity. There was no observed difference between doses. Without wishing to be held to any particular theory, the lack of difference between doses may be due to having crossed the threshold of drug uptake and concentration required in the mitochondria for a response.
- Human H460 NSCLC cells were obtained from American Type Cell Culture (ATCC) (catalog no. HTB-177, Manassas, VA). These cells tested negative for viral contamination using the Mouse Antibody Production (MAP) test, performed by Charles River Labs Molecular Division, upon the receipt of the tumor cells from ATCC.
- the tumor cells were maintained at 37 ° C in a humidified 5% CO2 atmosphere in T225 tissue culture flasks containing 50 mL of Roswell Park Memorial Institute (RPMI)-l640 solution with 10% Fetal Bovine Serum (FBS) and 2 mM L-glutamine. Cells were split at a ratio of 1: 10 every 2-3 days by trypsinization and resuspended in fresh medium in a new flask. Cells were harvested for experiments in the same way at 70-90% confluency.
- RPMI Roswell Park Memorial Institute
- FBS Fetal Bovine Serum
- CDl-Nu/Nu female mice ⁇ 4-6 weeks old were obtained from Charles River Laboratories. Mice were housed 5 to a cage in a micro-isolator room in the Department of Animal Laboratory Research of New York State University (SUNY) at Stony Brook. Light- dark cycles were 12 h each daily, with light from 7 a.m. to 7 p.m. Food (Purina Rodent Chow) and water (distilled sterile-filtered water, pH 7) were provided ad libitum. Protocols and procedures were according to the rules of and approved by the SUNY Institutional Animal Care and Use Committee (IACUC).
- IACUC Institutional Animal Care and Use Committee
- mice were inoculated subcutaneously (SC) in the right flank with 2xl0 6 human H460 NSCLC or BxPC3 pancreatic cancer cells that were suspended in 0.1 mL of Dulbeco’s Phosphate Buffered Salt (PBS) solution using a 1 cc syringe with a 27-5/8 gauge needle.
- PBS Phosphate Buffered Salt
- Tumor dimensions were measured daily before, during and after treatment (using Vernier calipers) and the tumor volume was calculated using the prolate ellipsoid formula: (length x width 2 )/2.
- Treatment with test or control articles began 8 days post tumor cell implantation when the tumor was approximately 300 mm 3 .
- Solid amorphous dispersion formulations of 6,8-bis-benzylthio-octanoic acid (API) were prepared by mixing the API 1:4 with one of the following polymers: Eudragit L100, poly(vinylpyrrolidone) viscosity grade K30 (PVP K30), hydroxypropyl methyl cellulose (HPMC), cellulose acetate phthalate (CAP), or hydroxypropyl methylcellulose acetate succinate (HPMCAS-M), and spray drying from methanol or acetone using a small-scale Bend Lab Dryer with 35 kg/hr drying gas flow rate capacity (BLD-35). Conditions, yields, and residual solvent levels of two representative spray dried dispersion (SDD) formulations (75 g each) are presented in the following table.
- SDD spray dried dispersion
- SEM Scanning electron microscopy
- X-ray diffraction is typically sensitive to the presence of crystalline material with an LOD of about 1% of the sample mass. No crystallinity was detected by PXRD for either SDD formulation.
- Diffractograms in comparison to crystalline 6,8-bis-benzylthio-octanoic acid API can be found in Figure 6, wherein the top diffractogram is the Eudragit L100 formulation, the middle diffractogram is the HPMCAS-M formulation, and the bottom diffractogram is crystalline 6,8-bis-benzylthio-octanoic acid.
- Example 6 Emulsion Oral Formulations of 6,8-Bis-benzylthio-octanoic Acid
- a 6,8-bis-benzylthio-octanoic acid solution was prepared by the steps of (a) providing a 50 mg/mL solution of 6,8-bis-benzylthio-octanoic acid in 1 M aqueous triethanolamine, and (b) diluting the 50 mg/mL solution with 5% aqueous dextrose to a concentration of 5 mg/mL.
- the resulting 5 mg/mL solution is identified as“7A” in Example 8 below.
- a suspension vehicle was prepared by the steps of: (a) combining tris buffer (48 mg) and HPMCAS-HF (20 mg) in 14 mL of distilled water, (b) adjusting the pH to 7.4 with dilute sodium hydroxide to dissolve the HPMCAS-HF, (c) heating the resulting solution to approximately 90°C, (d) adding Methocel A4M Premium (100 mg) to the hot solution, (e) stirring the mixture vigorously to suspend the undissolved Methocel A4M, (f) cooling and stirring the mixture with an ice bath until the Methocel A4M dissolves (approximately 10 minutes), (g) diluting the solution with distilled/deionized water to bring the total volume to 20 mL, and (h) adjusting the pH to 7.4 with dilute acetic acid or dilute sodium hydroxide to provide the suspension vehicle.
- Suspensions of the spray-dried formulations of Example 5 were prepared by adding 400 mg of the respective SDD formulation to a mortar, slowly adding 4 mL of the suspension vehicle (mixing thoroughly with a pestle after each small addition to uniformly disperse), and then transferring to a flask and stirring for one minute prior to administration.
- the resulting suspension of the Eudragit L100 SDD formulation (20 mg/mL 6,8-bis-benzylthio-octanoic acid) is identified as“7B” in Example 8 below.
- the resulting suspension of the HPMCAS-M SDD formulation (20 mg/mL 6,8-bis-benzylthio-octanoic acid) is identified as“7C” in
- a 20 mg/mL suspension of 6,8-bis-benzylthio-octanoic acid was prepared by adding 80 mg 6,8-bis-benzylthio-octanoic acid to a mortar, slowly adding 4 mL of the suspension vehicle (mixing thoroughly with a pestle after each small addition to uniformly disperse), and then transferring to a flask and stirring for one minute prior to administration.
- mice Six groups of 16 BALB/c nude mice (8 males and 8 females) per group were administered 6,8-bis-benzylthio-octanoic acid in six different ways: (1) 5 pL/g IV injection (tail vein) of the triethanolamine/dextrose aqueous solution of Example 7 (25 mg/kg; 5 mL/kg; Ex.
- Example 7A (2) 5 pL/g IP injection of the triethanolamine/dextrose aqueous solution of Example 7 (25 mg/kg; 5 mL/kg; 7 A); (3) 5 pL/g oral administration of the Eudragit L100 SDD suspension of Example 7 (100 mg/kg; 5 mL/kg; 7B); (4) 5 pL/g oral administration of the HPMCAS-M SDD suspension of Example 7 (100 mg/kg; 5 mL/kg; 7C); (5) 5 pL/g oral administration of the 20 mg/mL 6,8-bis-benzylthio-octanoic acid suspension of Example 7 (100 mg/kg; 5 mL/kg; 7D); or (6) 10 pL/g oral administration of the 10 mg/mL SOLUTOL solution of Example 7 (100 mg/kg; 10 mL/kg; 7E).
Landscapes
- Health & Medical Sciences (AREA)
- Chemical & Material Sciences (AREA)
- Life Sciences & Earth Sciences (AREA)
- Veterinary Medicine (AREA)
- Medicinal Chemistry (AREA)
- Pharmacology & Pharmacy (AREA)
- Animal Behavior & Ethology (AREA)
- General Health & Medical Sciences (AREA)
- Public Health (AREA)
- Epidemiology (AREA)
- Chemical Kinetics & Catalysis (AREA)
- General Chemical & Material Sciences (AREA)
- Engineering & Computer Science (AREA)
- Nuclear Medicine, Radiotherapy & Molecular Imaging (AREA)
- Organic Chemistry (AREA)
- Bioinformatics & Cheminformatics (AREA)
- Oil, Petroleum & Natural Gas (AREA)
- Dermatology (AREA)
- Nutrition Science (AREA)
- Physiology (AREA)
- Inorganic Chemistry (AREA)
- Oncology (AREA)
- Proteomics, Peptides & Aminoacids (AREA)
- Dispersion Chemistry (AREA)
- Biochemistry (AREA)
- Molecular Biology (AREA)
- Pharmaceuticals Containing Other Organic And Inorganic Compounds (AREA)
- Medicinal Preparation (AREA)
- Acyclic And Carbocyclic Compounds In Medicinal Compositions (AREA)
Abstract
Description
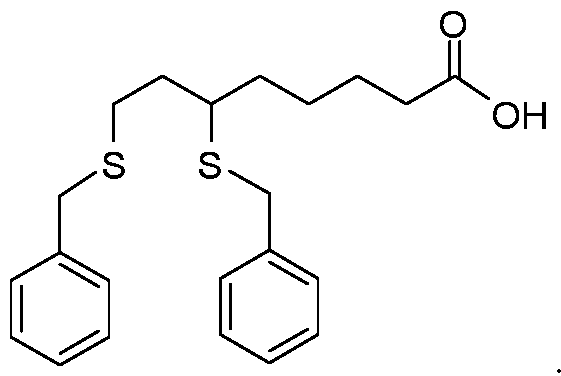

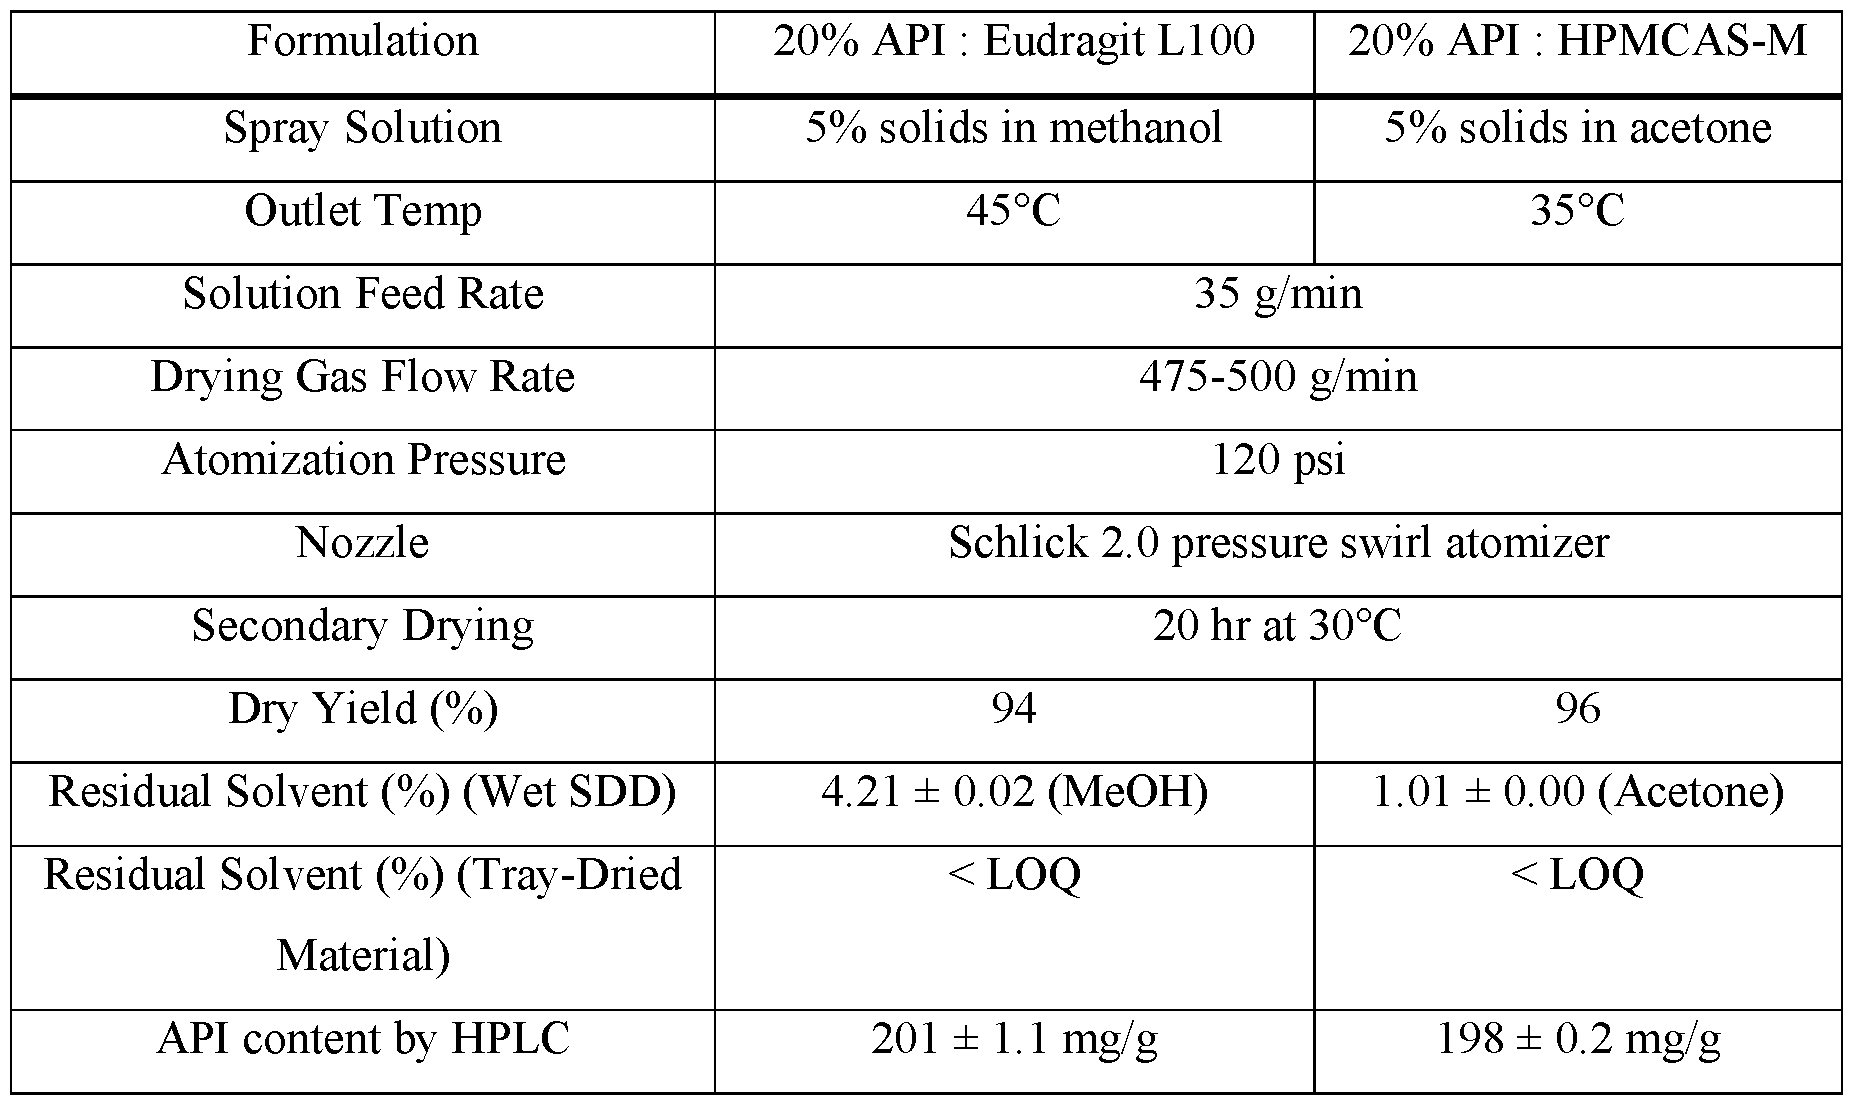

Claims
Priority Applications (5)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| CA3097380A CA3097380A1 (en) | 2018-04-16 | 2019-04-16 | Therapeutic methods and compositions for treating prostate cancer using 6,8-bis-benzylthio-octanoic acid |
| EP19789534.5A EP3781148A4 (en) | 2018-04-16 | 2019-04-16 | Therapeutic methods and compositions for treating prostate cancer using 6,8-bis-benzylthio-octanoic acid |
| KR1020207032586A KR20210023813A (en) | 2018-04-16 | 2019-04-16 | Treatment methods and compositions for treating prostate cancer using 6,8-bis-benzylthio-octanoic acid |
| US16/980,885 US20210000778A1 (en) | 2018-04-16 | 2019-04-16 | Therapeutic methods and compositions for treating prostate cancer using 6,8-bis-benzylthio-octanoic acid |
| JP2020556838A JP2021521222A (en) | 2018-04-16 | 2019-04-16 | Therapeutic methods and compositions for treating prostate cancer with 6,8-bis-benzylthio-octanoic acid |
Applications Claiming Priority (4)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| US201862658238P | 2018-04-16 | 2018-04-16 | |
| US62/658,238 | 2018-04-16 | ||
| US201862782938P | 2018-12-20 | 2018-12-20 | |
| US62/782,938 | 2018-12-20 |
Publications (1)
| Publication Number | Publication Date |
|---|---|
| WO2019204251A1 true WO2019204251A1 (en) | 2019-10-24 |
Family
ID=68239207
Family Applications (1)
| Application Number | Title | Priority Date | Filing Date |
|---|---|---|---|
| PCT/US2019/027597 Ceased WO2019204251A1 (en) | 2018-04-16 | 2019-04-16 | Therapeutic methods and compositions for treating prostate cancer using 6,8-bis-benzylthio-octanoic acid |
Country Status (6)
| Country | Link |
|---|---|
| US (1) | US20210000778A1 (en) |
| EP (1) | EP3781148A4 (en) |
| JP (1) | JP2021521222A (en) |
| KR (1) | KR20210023813A (en) |
| CA (1) | CA3097380A1 (en) |
| WO (1) | WO2019204251A1 (en) |
Cited By (2)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| WO2020132401A1 (en) * | 2018-12-20 | 2020-06-25 | Rafael Pharmaceuticals, Inc. | Oral therapy using 6,8-bis-benzylthio-octanoic acid |
| US20220265584A1 (en) * | 2019-07-11 | 2022-08-25 | Emory University | Combination Therapies for Managing Cancer |
Families Citing this family (3)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| US12297416B2 (en) | 2017-07-12 | 2025-05-13 | Deka Products Limited Partneship | System and method for transferring tissue |
| US10570362B2 (en) | 2017-07-12 | 2020-02-25 | Deka Products Limited Partnership | System and method for transferring tissue |
| AU2024204522B1 (en) | 2023-03-09 | 2024-08-22 | Minneamrita Therapeutics Llc | Drug combination for treatment of gastric cancer |
Citations (3)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| WO2011050261A1 (en) * | 2009-10-23 | 2011-04-28 | Cornerstone Pharmaceuticals, Inc. | Pharmaceutical formulations containing lipoic acid derivatives |
| US20130287791A1 (en) * | 2012-04-25 | 2013-10-31 | C. Wilson Xu | Modulation of histone h2b monoubiquitination and treatment of cancer |
| WO2018031407A1 (en) * | 2016-08-07 | 2018-02-15 | The Wistar Institute Of Anatomy And Biology | Methods of detecting and treating a tumor expressing pt346 pdk1 |
-
2019
- 2019-04-16 CA CA3097380A patent/CA3097380A1/en not_active Abandoned
- 2019-04-16 US US16/980,885 patent/US20210000778A1/en not_active Abandoned
- 2019-04-16 EP EP19789534.5A patent/EP3781148A4/en not_active Withdrawn
- 2019-04-16 WO PCT/US2019/027597 patent/WO2019204251A1/en not_active Ceased
- 2019-04-16 KR KR1020207032586A patent/KR20210023813A/en not_active Withdrawn
- 2019-04-16 JP JP2020556838A patent/JP2021521222A/en active Pending
Patent Citations (3)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| WO2011050261A1 (en) * | 2009-10-23 | 2011-04-28 | Cornerstone Pharmaceuticals, Inc. | Pharmaceutical formulations containing lipoic acid derivatives |
| US20130287791A1 (en) * | 2012-04-25 | 2013-10-31 | C. Wilson Xu | Modulation of histone h2b monoubiquitination and treatment of cancer |
| WO2018031407A1 (en) * | 2016-08-07 | 2018-02-15 | The Wistar Institute Of Anatomy And Biology | Methods of detecting and treating a tumor expressing pt346 pdk1 |
Non-Patent Citations (3)
| Title |
|---|
| LI, W ET AL.: "CD 44 regulates prostate cancer proliferation, invasion and migration via PDK1 and PFKB4", ONCOTARGET, vol. 8, 2017, pages 65143 - 65151, XP055649900 * |
| SAI, KKS ET AL.: "Abstract 4861: Initial preclinical evaluation of a novel inhibitor of mitochondrial metabolism against prostate cancer", CANCER RESEARCH, vol. 78, no. 13, July 2018 (2018-07-01), XP009516677 * |
| See also references of EP3781148A4 * |
Cited By (3)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| WO2020132401A1 (en) * | 2018-12-20 | 2020-06-25 | Rafael Pharmaceuticals, Inc. | Oral therapy using 6,8-bis-benzylthio-octanoic acid |
| US20220265584A1 (en) * | 2019-07-11 | 2022-08-25 | Emory University | Combination Therapies for Managing Cancer |
| US12377063B2 (en) * | 2019-07-11 | 2025-08-05 | Emory University | Combination therapies for managing cancer |
Also Published As
| Publication number | Publication date |
|---|---|
| US20210000778A1 (en) | 2021-01-07 |
| KR20210023813A (en) | 2021-03-04 |
| CA3097380A1 (en) | 2019-10-24 |
| EP3781148A4 (en) | 2022-03-02 |
| JP2021521222A (en) | 2021-08-26 |
| EP3781148A1 (en) | 2021-02-24 |
Similar Documents
| Publication | Publication Date | Title |
|---|---|---|
| US20210000778A1 (en) | Therapeutic methods and compositions for treating prostate cancer using 6,8-bis-benzylthio-octanoic acid | |
| US9610251B2 (en) | Pharmaceutical compositions for substituted quinazolinones | |
| US11622950B2 (en) | Therapeutic methods and compositions for treating pancreatic cancer using 6,8-bis-benzylthio-octanoic acid | |
| US20220054445A1 (en) | Therapeutic methods and compositions for treating cancer using 6,8-bis-benzylthio-octanoic acid and an autophagy inhibitor | |
| DK2528598T3 (en) | RELATIONS TO suppress PERIPHERAL NERVE DISORDER INDUCED BY AN ANTI-CANCER AGENT | |
| KR20180006422A (en) | Pharmaceutical compositions and uses thereof | |
| CA3121929A1 (en) | Therapeutic methods and compositions for treating cancer using 6,8-bis-benzylthio-octanoic acid and an autophagy inhibitor | |
| US20220040133A1 (en) | Oral therapy using 6,8-bis-benzylthio-octanoic acid | |
| US20080161404A1 (en) | Bicalutamide for Delivering Increasing Steady State Plasma Levels | |
| US20220331279A1 (en) | Therapeutic methods and compositions for treating cancer using 6,8-bis-benzylthio-octanoic acid and a glutaminase inhibitor | |
| US20240415791A1 (en) | Therapeutic methods and compositions for treating cancer using devimistat and a fatty acid oxidation inhibitor, a tyrosine kinase inhibitor, a glutaminase inhibitor, and/or a glycolysis inhibitor | |
| WO2021034631A1 (en) | Methods and pharmaceutical compositions containing 4,6-bis(benzylthio)hexanoic acid for treating cancer |
Legal Events
| Date | Code | Title | Description |
|---|---|---|---|
| 121 | Ep: the epo has been informed by wipo that ep was designated in this application |
Ref document number: 19789534 Country of ref document: EP Kind code of ref document: A1 |
|
| ENP | Entry into the national phase |
Ref document number: 2020556838 Country of ref document: JP Kind code of ref document: A |
|
| ENP | Entry into the national phase |
Ref document number: 3097380 Country of ref document: CA |
|
| NENP | Non-entry into the national phase |
Ref country code: DE |
|
| ENP | Entry into the national phase |
Ref document number: 20207032586 Country of ref document: KR Kind code of ref document: A |
|
| ENP | Entry into the national phase |
Ref document number: 2019789534 Country of ref document: EP Effective date: 20201116 |