EP0000745B1 - Salt of an optically active isomer of phenylglycine and an optically active isomer of 2-amino butanol and process for its preparation - Google Patents
Salt of an optically active isomer of phenylglycine and an optically active isomer of 2-amino butanol and process for its preparation Download PDFInfo
- Publication number
- EP0000745B1 EP0000745B1 EP78100531A EP78100531A EP0000745B1 EP 0000745 B1 EP0000745 B1 EP 0000745B1 EP 78100531 A EP78100531 A EP 78100531A EP 78100531 A EP78100531 A EP 78100531A EP 0000745 B1 EP0000745 B1 EP 0000745B1
- Authority
- EP
- European Patent Office
- Prior art keywords
- alkoxy
- salt
- optionally substituted
- optically active
- carbonyl
- Prior art date
- Legal status (The legal status is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the status listed.)
- Expired
Links
- 150000003839 salts Chemical class 0.000 title claims description 16
- JCBPETKZIGVZRE-UHFFFAOYSA-N 2-aminobutan-1-ol Chemical compound CCC(N)CO JCBPETKZIGVZRE-UHFFFAOYSA-N 0.000 title claims description 8
- 238000000034 method Methods 0.000 title claims description 8
- 238000002360 preparation method Methods 0.000 title claims description 3
- ZGUNAGUHMKGQNY-ZETCQYMHSA-N L-alpha-phenylglycine zwitterion Chemical compound OC(=O)[C@@H](N)C1=CC=CC=C1 ZGUNAGUHMKGQNY-ZETCQYMHSA-N 0.000 title 1
- -1 benzyliden Chemical group 0.000 claims description 16
- 230000003287 optical effect Effects 0.000 claims description 9
- 150000001875 compounds Chemical class 0.000 claims description 8
- 125000000217 alkyl group Chemical group 0.000 claims description 7
- 125000003545 alkoxy group Chemical group 0.000 claims description 6
- 125000000449 nitro group Chemical group [O-][N+](*)=O 0.000 claims description 6
- 229910052799 carbon Inorganic materials 0.000 claims description 4
- 125000004432 carbon atom Chemical group C* 0.000 claims description 4
- 125000005843 halogen group Chemical group 0.000 claims description 4
- 125000005076 adamantyloxycarbonyl group Chemical group C12(CC3CC(CC(C1)C3)C2)OC(=O)* 0.000 claims description 2
- 125000003435 aroyl group Chemical group 0.000 claims description 2
- 125000002102 aryl alkyloxo group Chemical group 0.000 claims description 2
- 125000004104 aryloxy group Chemical group 0.000 claims description 2
- 125000001584 benzyloxycarbonyl group Chemical group C(=O)(OCC1=CC=CC=C1)* 0.000 claims description 2
- 125000002915 carbonyl group Chemical group [*:2]C([*:1])=O 0.000 claims description 2
- GPAYUJZHTULNBE-UHFFFAOYSA-N diphenylphosphine Chemical compound C=1C=CC=CC=1PC1=CC=CC=C1 GPAYUJZHTULNBE-UHFFFAOYSA-N 0.000 claims description 2
- 238000001914 filtration Methods 0.000 claims description 2
- UYWQUFXKFGHYNT-UHFFFAOYSA-N phenylmethyl ester of formic acid Natural products O=COCC1=CC=CC=C1 UYWQUFXKFGHYNT-UHFFFAOYSA-N 0.000 claims description 2
- 125000001557 phthalyl group Chemical group C(=O)(O)C1=C(C(=O)*)C=CC=C1 0.000 claims description 2
- 125000002088 tosyl group Chemical group [H]C1=C([H])C(=C([H])C([H])=C1C([H])([H])[H])S(*)(=O)=O 0.000 claims description 2
- 125000002221 trityl group Chemical group [H]C1=C([H])C([H])=C([H])C([H])=C1C([*])(C1=C(C(=C(C(=C1[H])[H])[H])[H])[H])C1=C([H])C([H])=C([H])C([H])=C1[H] 0.000 claims description 2
- 229910052736 halogen Inorganic materials 0.000 claims 3
- QVGXLLKOCUKJST-UHFFFAOYSA-N atomic oxygen Chemical compound [O] QVGXLLKOCUKJST-UHFFFAOYSA-N 0.000 claims 2
- 150000001721 carbon Chemical group 0.000 claims 2
- 150000002367 halogens Chemical group 0.000 claims 2
- 229910052760 oxygen Inorganic materials 0.000 claims 2
- 239000001301 oxygen Substances 0.000 claims 2
- 125000000229 (C1-C4)alkoxy group Chemical group 0.000 claims 1
- 125000004423 acyloxy group Chemical group 0.000 claims 1
- 125000005194 alkoxycarbonyloxy group Chemical group 0.000 claims 1
- 125000000026 trimethylsilyl group Chemical group [H]C([H])([H])[Si]([*])(C([H])([H])[H])C([H])([H])[H] 0.000 claims 1
- OKKJLVBELUTLKV-UHFFFAOYSA-N Methanol Chemical compound OC OKKJLVBELUTLKV-UHFFFAOYSA-N 0.000 description 9
- LFQSCWFLJHTTHZ-UHFFFAOYSA-N Ethanol Chemical compound CCO LFQSCWFLJHTTHZ-UHFFFAOYSA-N 0.000 description 7
- 150000001413 amino acids Chemical class 0.000 description 6
- 150000001412 amines Chemical class 0.000 description 4
- 238000000926 separation method Methods 0.000 description 3
- KAGSUJOTNBINKR-UHFFFAOYSA-N 2-acetamido-2-(4-methoxyphenyl)acetic acid Chemical compound COC1=CC=C(C(NC(C)=O)C(O)=O)C=C1 KAGSUJOTNBINKR-UHFFFAOYSA-N 0.000 description 2
- KFZMGEQAYNKOFK-UHFFFAOYSA-N Isopropanol Chemical compound CC(C)O KFZMGEQAYNKOFK-UHFFFAOYSA-N 0.000 description 2
- WFRVJRZGHCTEGP-UHFFFAOYSA-N acetyl 2-acetamido-2-(4-hydroxyphenyl)acetate Chemical compound C(C)(=O)NC(C(=O)OC(C)=O)C1=CC=C(C=C1)O WFRVJRZGHCTEGP-UHFFFAOYSA-N 0.000 description 2
- 125000003277 amino group Chemical group 0.000 description 2
- 125000004430 oxygen atom Chemical group O* 0.000 description 2
- 239000000725 suspension Substances 0.000 description 2
- SANDMWGTFWJWOL-UHFFFAOYSA-N 2-(methoxyamino)-2-phenylacetic acid Chemical compound CONC(C(O)=O)C1=CC=CC=C1 SANDMWGTFWJWOL-UHFFFAOYSA-N 0.000 description 1
- 229930186147 Cephalosporin Natural products 0.000 description 1
- JVVXZOOGOGPDRZ-SLFFLAALSA-N [(1R,4aS,10aR)-1,4a-dimethyl-7-propan-2-yl-2,3,4,9,10,10a-hexahydrophenanthren-1-yl]methanamine Chemical compound NC[C@]1(C)CCC[C@]2(C)C3=CC=C(C(C)C)C=C3CC[C@H]21 JVVXZOOGOGPDRZ-SLFFLAALSA-N 0.000 description 1
- 150000003862 amino acid derivatives Chemical class 0.000 description 1
- 229960003022 amoxicillin Drugs 0.000 description 1
- LSQZJLSUYDQPKJ-NJBDSQKTSA-N amoxicillin Chemical compound C1([C@@H](N)C(=O)N[C@H]2[C@H]3SC([C@@H](N3C2=O)C(O)=O)(C)C)=CC=C(O)C=C1 LSQZJLSUYDQPKJ-NJBDSQKTSA-N 0.000 description 1
- 229960000723 ampicillin Drugs 0.000 description 1
- AVKUERGKIZMTKX-NJBDSQKTSA-N ampicillin Chemical compound C1([C@@H](N)C(=O)N[C@H]2[C@H]3SC([C@@H](N3C2=O)C(O)=O)(C)C)=CC=CC=C1 AVKUERGKIZMTKX-NJBDSQKTSA-N 0.000 description 1
- 239000003242 anti bacterial agent Substances 0.000 description 1
- 229940088710 antibiotic agent Drugs 0.000 description 1
- 229940124587 cephalosporin Drugs 0.000 description 1
- 150000001780 cephalosporins Chemical class 0.000 description 1
- KMPWYEUPVWOPIM-KODHJQJWSA-N cinchonidine Chemical compound C1=CC=C2C([C@H]([C@H]3[N@]4CC[C@H]([C@H](C4)C=C)C3)O)=CC=NC2=C1 KMPWYEUPVWOPIM-KODHJQJWSA-N 0.000 description 1
- KMPWYEUPVWOPIM-UHFFFAOYSA-N cinchonidine Natural products C1=CC=C2C(C(C3N4CCC(C(C4)C=C)C3)O)=CC=NC2=C1 KMPWYEUPVWOPIM-UHFFFAOYSA-N 0.000 description 1
- 239000012043 crude product Substances 0.000 description 1
- 238000004090 dissolution Methods 0.000 description 1
- 238000004519 manufacturing process Methods 0.000 description 1
- LSQZJLSUYDQPKJ-UHFFFAOYSA-N p-Hydroxyampicillin Natural products O=C1N2C(C(O)=O)C(C)(C)SC2C1NC(=O)C(N)C1=CC=C(O)C=C1 LSQZJLSUYDQPKJ-UHFFFAOYSA-N 0.000 description 1
- 125000001997 phenyl group Chemical group [H]C1=C([H])C([H])=C(*)C([H])=C1[H] 0.000 description 1
- 230000001376 precipitating effect Effects 0.000 description 1
- 238000001556 precipitation Methods 0.000 description 1
- 239000011541 reaction mixture Substances 0.000 description 1
- 238000011084 recovery Methods 0.000 description 1
- 238000001953 recrystallisation Methods 0.000 description 1
- 239000007858 starting material Substances 0.000 description 1
- 238000003756 stirring Methods 0.000 description 1
- QNLQKURWPIJSJS-UHFFFAOYSA-N trimethylsilylphosphane Chemical compound C[Si](C)(C)P QNLQKURWPIJSJS-UHFFFAOYSA-N 0.000 description 1
- XLYOFNOQVPJJNP-UHFFFAOYSA-N water Substances O XLYOFNOQVPJJNP-UHFFFAOYSA-N 0.000 description 1
Classifications
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07B—GENERAL METHODS OF ORGANIC CHEMISTRY; APPARATUS THEREFOR
- C07B57/00—Separation of optically-active compounds
Definitions
- the present invention relates to the separation into optical antipodes of compounds of the formula I. wherein XH, OH, C i --C 4- alkoxy, aryloxy, C 1 -C 4 -acyloxy, aralkyloxy, the optionally. is substituted with a maximum of 3 halogen atoms or nitro groups and contains a maximum of 4 carbon atoms in the alkyl radical, tert-alkoxy with a tertiary carbon atom bonded to the oxygen atom, aikoxycarbonyloxy or picolyloxycarbonyloxy and Y is aliphatic acyl, the counter.
- benzylidene may also be substituted with up to 3-halogen atoms or may contain further carbonyl groups, optionally halogen-substituted aroyl, or alkyl, alkoxy, nitro, phthalyl, trityl, optionally alkyl-, alkoxy- or halogen-substituted benzylidene, acetylisopropylidene, benzoylisopropylidene, 5,5-dimethyl-3-oxo-cyclohexen-1-yl, trichloroethyloxycarbonyl, benzyloxycarbonyl, optionally alkyl, alkoxy or halogen-substituted or substituted with a nitro group or ⁇ B (OH) 2 , tert-alkoxy with a tertiary carbon atom, which is attached to the oxygen atom, means adamantyloxycarbonyl, diphenylisopropyloxy
- optically active compounds of the formula mentioned are used as starting material for the production of semi-synthetic antibiotics of the cephalosporin or penicellin type, for example ampicillin or amoxycillin.
- the invention relates to a new process for the preparation of the optically active compounds of the above formula using optically active 2-aminobutanol.
- optically active natural amines such as cinchonidine or dehydroabiethylamine (cf. FR-A 2107926)
- these natural amines are very expensive and the separation yields are low.
- recovery of these natural amines for further use is limited because some of these amines are destroyed during the separation process, which is carried out at high temperatures.
- AT-PS 334 345 salts from optical isomers of N-acyl derivatives of dimethoxyphenylalanine and 2-aminobutanol are described. The yields are also low.
- the amino group of the amino acid or one of its derivatives is substituted in order to increase the acidity of the amino acid.
- These derivatives of the amino acids form the corresponding salts with 2-aminobutanol more easily than the amino acids themselves.
- the salts are generally prepared in water or a lower alkanol, preferably methanol, ethanol or isopropanol, one of the two salts precipitating from a racemic base Compound of formula I and optically active 2-aminobutanol can be formed.
- the method according to the invention is superior to known methods in that the op . table purity is higher than 99% and higher yields are obtained.
- amino acid derivatives are converted into the amino acids or into an amino acid which is either still substituted on the amino group or on the phenyl ring.
- the invention is illustrated by the following examples.
Landscapes
- Chemical & Material Sciences (AREA)
- Organic Chemistry (AREA)
- Organic Low-Molecular-Weight Compounds And Preparation Thereof (AREA)
Description
Die vorliegende Erfindung betrifft die Trennung in optische Antipoden von Verbindungen der Formel I

Die optisch aktiven Verbindungen der genannten Formel werden als Ausgangsmaterial für die Produktion von halbsynthetischen Antibiotika des Cephalosporin oder Penicellin-Typs, beispielsweise Ampicillin oder Amoxycillin eingesetzt. Die Erfindung betrifft ein neues Verfahren zur Herstellung der optisch aktiven Verbindungen der obigen Formel unter Einsatz von optisch aktivem 2-Aminobutanol.The optically active compounds of the formula mentioned are used as starting material for the production of semi-synthetic antibiotics of the cephalosporin or penicellin type, for example ampicillin or amoxycillin. The invention relates to a new process for the preparation of the optically active compounds of the above formula using optically active 2-aminobutanol.
In einem bekannten Verfahren zur Trennung von Verbindungen in optische Isomere der obigen Formel wurden optisch aktive natürliche Amine, wie Cinchonidin oder Dehydroabiethylamin (vgl. FR-A 2107926), eingesetzt. Doch diese natürlichen Amine sind sehr teuer, und die Trennungsausbeuten sind niedrig. Hinzu kommt noch der weitere Nachteil, daß eine Wiedergewinnung dieser natürlichen Amine zum weiteren Gebrauch begrenzt ist, weil ein Teil dieser Amine während des Trennungsverfahrens, das mit hohen Temperaturen durchgeführt wird, zerstört wird. In der AT-PS 334 345 werden Salze aus optischen Isomeren von N-Acylderivaten von Dimethoxyphenylalanin und 2-Aminobutanol beschrieben. Die Ausbeuten sind ebenfalls gering. Gemäß der Erfindung wird die Aminogruppe der Aminosäure oder eines ihrer Derivate substituiert, um die Acidität der Aminosäure zu erhöhen. Diese Derivate der Aminosäuren bilden mit 2-Aminobutanol leichter die entsprechenden Salze als die Aminosäuren selbst. Die Salze werden im allgemeinen in Wasser oder einem niederen Alkanol, vorzugsweise Methanol, Äthanol oder Isopropanol hergestellt, wobei eines der beiden Salze ausfällt, die ausgehend von einer racemischen Verbindung der Formel I und optisch aktivem 2-Aminobutanol gebildet werden können.In a known process for separating compounds into optical isomers of the above formula, optically active natural amines, such as cinchonidine or dehydroabiethylamine (cf. FR-A 2107926), have been used. However, these natural amines are very expensive and the separation yields are low. In addition, there is the further disadvantage that recovery of these natural amines for further use is limited because some of these amines are destroyed during the separation process, which is carried out at high temperatures. In AT-PS 334 345 salts from optical isomers of N-acyl derivatives of dimethoxyphenylalanine and 2-aminobutanol are described. The yields are also low. According to the invention, the amino group of the amino acid or one of its derivatives is substituted in order to increase the acidity of the amino acid. These derivatives of the amino acids form the corresponding salts with 2-aminobutanol more easily than the amino acids themselves. The salts are generally prepared in water or a lower alkanol, preferably methanol, ethanol or isopropanol, one of the two salts precipitating from a racemic base Compound of formula I and optically active 2-aminobutanol can be formed.
Das erfindungsgemäße Verfahren ist bekannten Verfahren darin überlegen, daß die op-. tische Reinheit höher als 99% liegt und höhere Ausbeuten erhalten werden.The method according to the invention is superior to known methods in that the op . table purity is higher than 99% and higher yields are obtained.
Gegebenenfalls werden die AminosäureDerivate in die Aminosäuren umgewandelt oder in eine Aminosäure, die entweder noch an der Aminogruppe oder am Phenylring substituiert ist. Die Erfindung wird durch die folgenden Beispiele erläutert.If appropriate, the amino acid derivatives are converted into the amino acids or into an amino acid which is either still substituted on the amino group or on the phenyl ring. The invention is illustrated by the following examples.
Salz von 1-N,O-Diacetyl-4-hydroxy-phenylglycin und 1-2-Aminobutanol.Salt of 1-N, O-diacetyl-4-hydroxy-phenylglycine and 1-2-aminobutanol.
Zu einer Suspension von dl-Diacetyl-4-hydroxy-phenylglycin (47,5 g) in 400 ml Äthanol, wurde 1-2-Aminobutanol (18 g) unter Rühren gegeben. Die Reaktionsmischung wurde langsam bis zur Auflösung erwärmt, worauf auf Raumtemperatur abgekühlt und 2 Stunden stehengelassen wurde, um die Ausfällung des Salzes zu vervollständigen. Das Salz wurde durch Filtrieren abgetrennt und ergab ein Rohprodukt (30 g). Durch Umkristallisieren aus 80 ml Äthanol wurde das reine Salz aus 1-N,O-Diacetyl-4-hydroxy-phenylglycin und 1-2-Aminobutanol erhalten.To a suspension of dl-diacetyl-4-hydroxy-phenylglycine (47.5 g) in 400 ml of ethanol was added 1-2-aminobutanol (18 g) with stirring. The reaction mixture was slowly warmed to dissolution, then cooled to room temperature and allowed to stand for 2 hours to complete the precipitation of the salt. The salt was separated by filtration and gave a crude product (30 g). The pure salt of 1-N, O-diacetyl-4-hydroxy-phenylglycine and 1-2-aminobutanol was obtained by recrystallization from 80 ml of ethanol.
Fp. 168-170° C und (∝)25 D = -126° (C=2, H2 0).Mp 168-170 ° C and (∝) 25 D = -126 ° (C = 2, H 2 0 ).
Das gemäß obiger Arbeitsweise erhaltene Salz wurde in bekannter Weise behandelt, um 1-4-Hydroxyphenylglycin mit einer optischen Reinheit ([a]25 = -159° (C=2, N-HCI) von mehr als 99% zu erhalten.The salt obtained in accordance with the above procedure was treated in a known manner in order to obtain 1-4-hydroxyphenylglycine with an optical purity ([a] 2 5 = -159 ° (C = 2, N-HCl) of more than 99%.
Salz von 1-N-Acetyl-4-methoxy-phenylglycin mit 1-2-Aminobutanol.Salt of 1-N-acetyl-4-methoxy-phenylglycine with 1-2-aminobutanol.
Zu einer Suspension von 1-N-Acetyl-4-methoxy-phenylglycin (4,46 g) in 15 ml abs. Methanol wurde 1-2-Aminobutanol (1,8g) gegeben. Es wurde wie in Beispiel 1 verfahren und ein rohes Salz (3,1 g) von 1-N-Acetyl-4-methoxy-phenylglycin mit 1-2-Aminobutanot erhalten. Das reine Salz wurde durch Um-. kristallisieren aus 5 ml abs. Methanol erhalten. 2,5 g (80%) 
Claims (2)
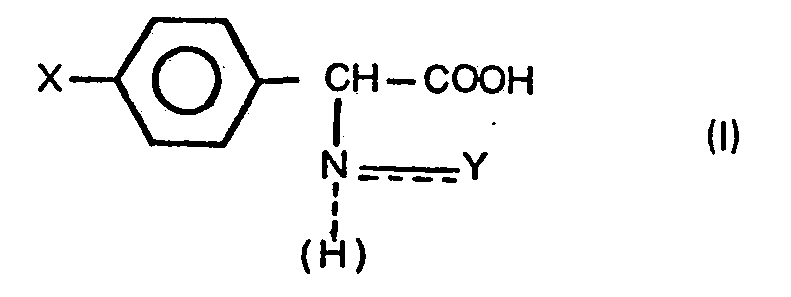
Applications Claiming Priority (2)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| DE2735834 | 1977-08-09 | ||
| DE19772735834 DE2735834A1 (en) | 1977-08-09 | 1977-08-09 | SEPARATION OF AMINO ACIDS INTO OPTICAL ANTIPODES |
Publications (2)
| Publication Number | Publication Date |
|---|---|
| EP0000745A1 EP0000745A1 (en) | 1979-02-21 |
| EP0000745B1 true EP0000745B1 (en) | 1981-07-15 |
Family
ID=6015982
Family Applications (1)
| Application Number | Title | Priority Date | Filing Date |
|---|---|---|---|
| EP78100531A Expired EP0000745B1 (en) | 1977-08-09 | 1978-07-28 | Salt of an optically active isomer of phenylglycine and an optically active isomer of 2-amino butanol and process for its preparation |
Country Status (14)
| Country | Link |
|---|---|
| US (1) | US4182899A (en) |
| EP (1) | EP0000745B1 (en) |
| JP (1) | JPS5430132A (en) |
| AT (1) | AT359997B (en) |
| BG (1) | BG35593A3 (en) |
| CA (1) | CA1110653A (en) |
| CS (1) | CS203200B2 (en) |
| DD (1) | DD137580A5 (en) |
| DE (2) | DE2735834A1 (en) |
| HU (1) | HU178248B (en) |
| IE (1) | IE47236B1 (en) |
| IL (1) | IL55293A (en) |
| IT (1) | IT1098014B (en) |
| SU (1) | SU913937A3 (en) |
Families Citing this family (7)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| HU193199B (en) * | 1984-03-01 | 1987-08-28 | Budapesti Mueszaki Egyetem | Process for preparing optically active alpha-amino-beta-phenyl-propionic acids |
| EP0174358A1 (en) * | 1984-03-01 | 1986-03-19 | ALKALOIDA VEGYéSZETI GYáR | Novel diastereomer salts of phenylalanine and n-acyl derivatives thereof and process for the separation of optically active phenylalanine and n-acyl derivatives thereof |
| JPH01155119A (en) * | 1987-12-14 | 1989-06-19 | Rinnai Corp | Combustion control device |
| IT1226903B (en) * | 1988-07-12 | 1991-02-21 | Mini Ricerca Scient Tecnolog | PROCESS FOR THE SYNTHESIS OF OPTICALLY ACTIVE AMINO ACIDS |
| US5583259A (en) * | 1991-02-08 | 1996-12-10 | Les Laboratoires Beecham S.A. | 2-(RO)-1-(R) ethylamines |
| FR2672593B1 (en) * | 1991-02-08 | 1994-05-20 | Beecham Sa Laboratoires | NEW COMPOUNDS USEFUL FOR THE RESOLUTION OF DL-N-ACETYL OR DL-N-HALOACETYL-HYDROXYPHENYLGLYCINE. |
| CN102887836B (en) * | 2011-07-18 | 2014-03-26 | 西南大学 | L-phenylglycine derivative and application thereof |
Family Cites Families (8)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| NL268416A (en) * | 1960-08-19 | |||
| US3869505A (en) * | 1970-09-24 | 1975-03-04 | Beecham Group Ltd | DL-2-(4-hydroxyphenyl)-glycine-N,O-diacetate and salts |
| US3904681A (en) * | 1971-03-30 | 1975-09-09 | Ciba Geigy Corp | Propionic acid |
| US3796748A (en) * | 1972-08-16 | 1974-03-12 | Bristol Myers Co | Dehydroabietylammonium d-(-)-2-chloroacetylamino-2-(p-hydroxyphenyl)-acetate |
| YU36484B (en) * | 1973-06-07 | 1984-02-29 | Pliva Pharm & Chem Works | Process for preparing l-(-)-2-methyl-3-(3,4-dihydroxyphenyl)-alanine |
| JPS5069039A (en) * | 1973-10-23 | 1975-06-09 | ||
| JPS50116434A (en) * | 1974-03-01 | 1975-09-11 | ||
| JPS5152154A (en) * | 1974-10-26 | 1976-05-08 | Sankyo Co | nn karubometokishifuenirugurishinno kogakubunkatsuho |
-
1977
- 1977-08-09 DE DE19772735834 patent/DE2735834A1/en not_active Withdrawn
-
1978
- 1978-07-28 EP EP78100531A patent/EP0000745B1/en not_active Expired
- 1978-07-28 DE DE7878100531T patent/DE2860839D1/en not_active Expired
- 1978-08-04 DD DD78207128A patent/DD137580A5/en unknown
- 1978-08-04 BG BG040601A patent/BG35593A3/en unknown
- 1978-08-07 IT IT26559/78A patent/IT1098014B/en active
- 1978-08-07 IL IL55293A patent/IL55293A/en unknown
- 1978-08-07 CS CS785160A patent/CS203200B2/en unknown
- 1978-08-07 US US05/931,529 patent/US4182899A/en not_active Expired - Lifetime
- 1978-08-07 HU HU78HO2093A patent/HU178248B/en unknown
- 1978-08-08 SU SU782645702A patent/SU913937A3/en active
- 1978-08-08 AT AT577278A patent/AT359997B/en not_active IP Right Cessation
- 1978-08-08 CA CA308,911A patent/CA1110653A/en not_active Expired
- 1978-08-08 IE IE1611/78A patent/IE47236B1/en unknown
- 1978-08-09 JP JP9628578A patent/JPS5430132A/en active Pending
Also Published As
| Publication number | Publication date |
|---|---|
| IE781611L (en) | 1979-02-09 |
| ATA577278A (en) | 1980-05-15 |
| HU178248B (en) | 1982-04-28 |
| IT1098014B (en) | 1985-08-31 |
| IL55293A0 (en) | 1978-10-31 |
| DE2860839D1 (en) | 1981-10-22 |
| IT7826559A0 (en) | 1978-08-07 |
| BG35593A3 (en) | 1984-05-15 |
| SU913937A3 (en) | 1982-03-15 |
| JPS5430132A (en) | 1979-03-06 |
| EP0000745A1 (en) | 1979-02-21 |
| AT359997B (en) | 1980-12-10 |
| CS203200B2 (en) | 1981-02-27 |
| IL55293A (en) | 1982-09-30 |
| US4182899A (en) | 1980-01-08 |
| IE47236B1 (en) | 1984-01-25 |
| DE2735834A1 (en) | 1979-02-22 |
| DD137580A5 (en) | 1979-09-12 |
| CA1110653A (en) | 1981-10-13 |
Similar Documents
| Publication | Publication Date | Title |
|---|---|---|
| WO1993010118A1 (en) | Method of separating the enantiomers of 5-methyl-tetrahydrofolic acid | |
| EP0625137B1 (en) | Method of separating racemates of verapamil | |
| EP0312726B1 (en) | Optically active salt of a substituted thiazolidine-4-carboxylic acid and a 3-chlor-2-hydroxypropyl trimethyl ammonium, their preparation and use | |
| EP0000745A1 (en) | Salt of an optically active isomer of phenylglycine and an optically active isomer of 2-amino butanol and process for its preparation | |
| EP0237902A2 (en) | Process for the preparation of the (-)-antipodes of (E)-1-cyclohexyl-4,4-dimethyl-3-hydroxy-2-(1,2,4-triazol-1-yl)-pent-1-enes | |
| DE2926847C2 (en) | ||
| DE2501957C2 (en) | Process for the production of optically active p-hydroxyphenylglycine | |
| EP0126934B1 (en) | Process for the preparation of 1,1-disubstituted thioureas | |
| DE69016922T2 (en) | Process for the preparation of 17-esters of 21-deoxyprednisolone. | |
| DE1518337A1 (en) | New X-ray contrast media and methods of making same | |
| DE2141765A1 (en) | 4-chloro-4-thiazolin-2-ones - with microbicidal properties | |
| DE69113365T2 (en) | Optically active morpholinooxobutanoic acid hydroxybinaphthalene derivative and its preparation. | |
| DE2150267C3 (en) | Process for the production of peptides | |
| DE1933187B2 (en) | Process for the preparation of 7-aminocephalosporanic acid | |
| EP0073052B1 (en) | Process for the resolution of the racemate s-(carboxymethyl)-(rs)-cysteine (a) | |
| EP0478721B1 (en) | New benzocycloalkene carboxylic acid and process for producing it | |
| EP0315097A2 (en) | Acyl labdane derivatives, process for their preparation and their use as medicines | |
| DE869964C (en) | Process for the preparation of choline derivatives | |
| DE1192209B (en) | Process for the preparation of esters of quaternary ammonium compounds | |
| DE2623076A1 (en) | ENAMINES AND PROCEDURES FOR THEIR PRODUCTION | |
| DE1445692A1 (en) | Process for the preparation of piperazine derivatives | |
| DE1057133B (en) | Process for the production of new, pharmacologically active 1,1-diphenyl-2-aminoalkanols | |
| DE2535693A1 (en) | NEW ESTERS OF PGF DEEP 2ALPHA, 15 (S) -15-METHYL-PGF DEEP 2ALPHA AND 15 (R) -15-METHYL-PGF DEEP 2ALPHA | |
| DE2154437A1 (en) | Process for the preparation of substituted alpha-naphthoxyacetamides | |
| CH649526A5 (en) | Process for the preparation of (Z)-1,2-diphenyl-1-[p-(2-dimethyl-aminoethoxy)phenyl]but-1-ene |
Legal Events
| Date | Code | Title | Description |
|---|---|---|---|
| PUAI | Public reference made under article 153(3) epc to a published international application that has entered the european phase |
Free format text: ORIGINAL CODE: 0009012 |
|
| AK | Designated contracting states |
Designated state(s): BE CH DE FR GB NL SE |
|
| 17P | Request for examination filed | ||
| GRAA | (expected) grant |
Free format text: ORIGINAL CODE: 0009210 |
|
| AK | Designated contracting states |
Designated state(s): BE CH DE FR GB NL SE |
|
| REF | Corresponds to: |
Ref document number: 2860839 Country of ref document: DE Date of ref document: 19811022 |
|
| PGFP | Annual fee paid to national office [announced via postgrant information from national office to epo] |
Ref country code: CH Payment date: 19840618 Year of fee payment: 7 |
|
| PGFP | Annual fee paid to national office [announced via postgrant information from national office to epo] |
Ref country code: FR Payment date: 19840621 Year of fee payment: 7 |
|
| PGFP | Annual fee paid to national office [announced via postgrant information from national office to epo] |
Ref country code: BE Payment date: 19840630 Year of fee payment: 7 |
|
| PGFP | Annual fee paid to national office [announced via postgrant information from national office to epo] |
Ref country code: DE Payment date: 19840810 Year of fee payment: 7 |
|
| PGFP | Annual fee paid to national office [announced via postgrant information from national office to epo] |
Ref country code: SE Payment date: 19840930 Year of fee payment: 7 |
|
| PGFP | Annual fee paid to national office [announced via postgrant information from national office to epo] |
Ref country code: NL Payment date: 19850731 Year of fee payment: 8 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: DE Effective date: 19860402 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: SE Effective date: 19860729 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: CH Effective date: 19860731 Ref country code: BE Effective date: 19860731 |
|
| BERE | Be: lapsed |
Owner name: HOECHST A.G. Effective date: 19860731 |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: NL Effective date: 19870201 |
|
| NLV4 | Nl: lapsed or anulled due to non-payment of the annual fee | ||
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: FR Free format text: LAPSE BECAUSE OF NON-PAYMENT OF DUE FEES Effective date: 19870331 |
|
| REG | Reference to a national code |
Ref country code: CH Ref legal event code: PL |
|
| GBPC | Gb: european patent ceased through non-payment of renewal fee | ||
| REG | Reference to a national code |
Ref country code: FR Ref legal event code: ST |
|
| PG25 | Lapsed in a contracting state [announced via postgrant information from national office to epo] |
Ref country code: GB Effective date: 19881117 |
|
| EUG | Se: european patent has lapsed |
Ref document number: 78100531.9 Effective date: 19870505 |
|
| PLBE | No opposition filed within time limit |
Free format text: ORIGINAL CODE: 0009261 |
|
| STAA | Information on the status of an ep patent application or granted ep patent |
Free format text: STATUS: NO OPPOSITION FILED WITHIN TIME LIMIT |