JP2012246231A - Method for producing heterocyclic compound - Google Patents
Method for producing heterocyclic compound Download PDFInfo
- Publication number
- JP2012246231A JP2012246231A JP2011117605A JP2011117605A JP2012246231A JP 2012246231 A JP2012246231 A JP 2012246231A JP 2011117605 A JP2011117605 A JP 2011117605A JP 2011117605 A JP2011117605 A JP 2011117605A JP 2012246231 A JP2012246231 A JP 2012246231A
- Authority
- JP
- Japan
- Prior art keywords
- group
- heterocyclic compound
- production method
- atom
- mol
- Prior art date
- Legal status (The legal status is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the status listed.)
- Granted
Links
- 238000004519 manufacturing process Methods 0.000 title claims abstract description 28
- 150000002391 heterocyclic compounds Chemical class 0.000 title claims abstract description 12
- 238000006243 chemical reaction Methods 0.000 claims abstract description 13
- 125000005843 halogen group Chemical group 0.000 claims abstract description 6
- XLYOFNOQVPJJNP-UHFFFAOYSA-N water Substances O XLYOFNOQVPJJNP-UHFFFAOYSA-N 0.000 claims description 17
- HEMHJVSKTPXQMS-UHFFFAOYSA-M Sodium hydroxide Chemical compound [OH-].[Na+] HEMHJVSKTPXQMS-UHFFFAOYSA-M 0.000 claims description 12
- 125000004435 hydrogen atom Chemical group [H]* 0.000 claims description 12
- BWHMMNNQKKPAPP-UHFFFAOYSA-L potassium carbonate Chemical compound [K+].[K+].[O-]C([O-])=O BWHMMNNQKKPAPP-UHFFFAOYSA-L 0.000 claims description 10
- 125000000217 alkyl group Chemical group 0.000 claims description 9
- XLOMVQKBTHCTTD-UHFFFAOYSA-N Zinc monoxide Chemical compound [Zn]=O XLOMVQKBTHCTTD-UHFFFAOYSA-N 0.000 claims description 8
- 125000003545 alkoxy group Chemical group 0.000 claims description 8
- 125000001309 chloro group Chemical group Cl* 0.000 claims description 8
- 150000001875 compounds Chemical class 0.000 claims description 8
- 125000001997 phenyl group Chemical group [H]C1=C([H])C([H])=C(*)C([H])=C1[H] 0.000 claims description 8
- 125000001424 substituent group Chemical group 0.000 claims description 8
- 229910052801 chlorine Inorganic materials 0.000 claims description 7
- 125000004093 cyano group Chemical group *C#N 0.000 claims description 7
- VTYYLEPIZMXCLO-UHFFFAOYSA-L Calcium carbonate Chemical compound [Ca+2].[O-]C([O-])=O VTYYLEPIZMXCLO-UHFFFAOYSA-L 0.000 claims description 6
- CDBYLPFSWZWCQE-UHFFFAOYSA-L Sodium Carbonate Chemical compound [Na+].[Na+].[O-]C([O-])=O CDBYLPFSWZWCQE-UHFFFAOYSA-L 0.000 claims description 6
- UIIMBOGNXHQVGW-UHFFFAOYSA-M Sodium bicarbonate Chemical compound [Na+].OC([O-])=O UIIMBOGNXHQVGW-UHFFFAOYSA-M 0.000 claims description 6
- 238000006467 substitution reaction Methods 0.000 claims description 6
- 239000003054 catalyst Substances 0.000 claims description 5
- 150000007529 inorganic bases Chemical class 0.000 claims description 5
- 229910000027 potassium carbonate Inorganic materials 0.000 claims description 5
- 230000001737 promoting effect Effects 0.000 claims description 5
- ZCYVEMRRCGMTRW-UHFFFAOYSA-N 7553-56-2 Chemical group [I] ZCYVEMRRCGMTRW-UHFFFAOYSA-N 0.000 claims description 4
- WKBOTKDWSSQWDR-UHFFFAOYSA-N Bromine atom Chemical group [Br] WKBOTKDWSSQWDR-UHFFFAOYSA-N 0.000 claims description 4
- 229910052740 iodine Inorganic materials 0.000 claims description 4
- 239000011787 zinc oxide Substances 0.000 claims description 4
- 125000006273 (C1-C3) alkyl group Chemical group 0.000 claims description 3
- 229910000019 calcium carbonate Inorganic materials 0.000 claims description 3
- GBRBMTNGQBKBQE-UHFFFAOYSA-L copper;diiodide Chemical compound I[Cu]I GBRBMTNGQBKBQE-UHFFFAOYSA-L 0.000 claims description 3
- 125000000449 nitro group Chemical group [O-][N+](*)=O 0.000 claims description 3
- 229910000030 sodium bicarbonate Inorganic materials 0.000 claims description 3
- 235000017557 sodium bicarbonate Nutrition 0.000 claims description 3
- 229910000029 sodium carbonate Inorganic materials 0.000 claims description 3
- 150000001420 substituted heterocyclic compounds Chemical class 0.000 claims description 3
- 125000002490 anilino group Chemical class [H]N(*)C1=C([H])C([H])=C([H])C([H])=C1[H] 0.000 claims 1
- -1 pyridine compound Chemical class 0.000 abstract description 18
- PAYRUJLWNCNPSJ-UHFFFAOYSA-N Aniline Chemical compound NC1=CC=CC=C1 PAYRUJLWNCNPSJ-UHFFFAOYSA-N 0.000 abstract description 10
- JUJWROOIHBZHMG-UHFFFAOYSA-N pyridine Substances C1=CC=NC=C1 JUJWROOIHBZHMG-UHFFFAOYSA-N 0.000 abstract description 8
- UMJSCPRVCHMLSP-UHFFFAOYSA-N pyridine Natural products COC1=CC=CN=C1 UMJSCPRVCHMLSP-UHFFFAOYSA-N 0.000 abstract description 8
- LSPMHHJCDSFAAY-UHFFFAOYSA-N 2,6-dichloro-4-methylpyridine-3-carbonitrile Chemical compound CC1=CC(Cl)=NC(Cl)=C1C#N LSPMHHJCDSFAAY-UHFFFAOYSA-N 0.000 abstract description 6
- 150000001448 anilines Chemical class 0.000 abstract description 6
- 238000000034 method Methods 0.000 abstract description 5
- 239000000126 substance Substances 0.000 abstract description 4
- VEXZGXHMUGYJMC-UHFFFAOYSA-N Hydrochloric acid Chemical compound Cl VEXZGXHMUGYJMC-UHFFFAOYSA-N 0.000 description 14
- 239000000203 mixture Substances 0.000 description 12
- 239000000047 product Substances 0.000 description 11
- 239000013078 crystal Substances 0.000 description 9
- 239000006227 byproduct Substances 0.000 description 8
- 238000001914 filtration Methods 0.000 description 7
- 150000003839 salts Chemical class 0.000 description 6
- 238000003756 stirring Methods 0.000 description 6
- 125000004191 (C1-C6) alkoxy group Chemical group 0.000 description 4
- KFZMGEQAYNKOFK-UHFFFAOYSA-N Isopropanol Chemical compound CC(C)O KFZMGEQAYNKOFK-UHFFFAOYSA-N 0.000 description 4
- 239000000706 filtrate Substances 0.000 description 4
- 125000002496 methyl group Chemical group [H]C([H])([H])* 0.000 description 4
- 230000007935 neutral effect Effects 0.000 description 4
- 239000002904 solvent Substances 0.000 description 4
- WEVYAHXRMPXWCK-UHFFFAOYSA-N Acetonitrile Chemical compound CC#N WEVYAHXRMPXWCK-UHFFFAOYSA-N 0.000 description 3
- 239000002994 raw material Substances 0.000 description 3
- 239000003153 chemical reaction reagent Substances 0.000 description 2
- 239000007810 chemical reaction solvent Substances 0.000 description 2
- 230000000052 comparative effect Effects 0.000 description 2
- 239000000975 dye Substances 0.000 description 2
- 125000001495 ethyl group Chemical group [H]C([H])([H])C([H])([H])* 0.000 description 2
- 238000004128 high performance liquid chromatography Methods 0.000 description 2
- 238000011141 high resolution liquid chromatography Methods 0.000 description 2
- 125000001449 isopropyl group Chemical group [H]C([H])([H])C([H])(*)C([H])([H])[H] 0.000 description 2
- 125000004123 n-propyl group Chemical group [H]C([H])([H])C([H])([H])C([H])([H])* 0.000 description 2
- 230000002194 synthesizing effect Effects 0.000 description 2
- 125000004169 (C1-C6) alkyl group Chemical group 0.000 description 1
- QTBSBXVTEAMEQO-UHFFFAOYSA-M Acetate Chemical compound CC([O-])=O QTBSBXVTEAMEQO-UHFFFAOYSA-M 0.000 description 1
- QGZKDVFQNNGYKY-UHFFFAOYSA-O Ammonium Chemical compound [NH4+] QGZKDVFQNNGYKY-UHFFFAOYSA-O 0.000 description 1
- BVKZGUZCCUSVTD-UHFFFAOYSA-L Carbonate Chemical compound [O-]C([O-])=O BVKZGUZCCUSVTD-UHFFFAOYSA-L 0.000 description 1
- NHNBFGGVMKEFGY-UHFFFAOYSA-N Nitrate Chemical compound [O-][N+]([O-])=O NHNBFGGVMKEFGY-UHFFFAOYSA-N 0.000 description 1
- QAOWNCQODCNURD-UHFFFAOYSA-L Sulfate Chemical compound [O-]S([O-])(=O)=O QAOWNCQODCNURD-UHFFFAOYSA-L 0.000 description 1
- 150000001450 anions Chemical class 0.000 description 1
- 230000015572 biosynthetic process Effects 0.000 description 1
- 125000000484 butyl group Chemical group [H]C([*])([H])C([H])([H])C([H])([H])C([H])([H])[H] 0.000 description 1
- 125000003178 carboxy group Chemical group [H]OC(*)=O 0.000 description 1
- 150000001768 cations Chemical class 0.000 description 1
- 239000003795 chemical substances by application Substances 0.000 description 1
- 238000002425 crystallisation Methods 0.000 description 1
- 230000008025 crystallization Effects 0.000 description 1
- 230000008034 disappearance Effects 0.000 description 1
- 239000000986 disperse dye Substances 0.000 description 1
- 239000012776 electronic material Substances 0.000 description 1
- 125000001301 ethoxy group Chemical group [H]C([H])([H])C([H])([H])O* 0.000 description 1
- 239000000835 fiber Substances 0.000 description 1
- 238000010438 heat treatment Methods 0.000 description 1
- 125000002887 hydroxy group Chemical group [H]O* 0.000 description 1
- 125000002510 isobutoxy group Chemical group [H]C([H])([H])C([H])(C([H])([H])[H])C([H])([H])O* 0.000 description 1
- 125000003253 isopropoxy group Chemical group [H]C([H])([H])C([H])(O*)C([H])([H])[H] 0.000 description 1
- 229910021645 metal ion Inorganic materials 0.000 description 1
- 125000000956 methoxy group Chemical group [H]C([H])([H])O* 0.000 description 1
- 125000006606 n-butoxy group Chemical group 0.000 description 1
- 125000004108 n-butyl group Chemical group [H]C([H])([H])C([H])([H])C([H])([H])C([H])([H])* 0.000 description 1
- 125000003136 n-heptyl group Chemical group [H]C([H])([H])C([H])([H])C([H])([H])C([H])([H])C([H])([H])C([H])([H])C([H])([H])* 0.000 description 1
- 125000001280 n-hexyl group Chemical group C(CCCCC)* 0.000 description 1
- 125000003935 n-pentoxy group Chemical group [H]C([H])([H])C([H])([H])C([H])([H])C([H])([H])C([H])([H])O* 0.000 description 1
- 125000003506 n-propoxy group Chemical group [H]C([H])([H])C([H])([H])C([H])([H])O* 0.000 description 1
- 239000003960 organic solvent Substances 0.000 description 1
- 239000000575 pesticide Substances 0.000 description 1
- NBIIXXVUZAFLBC-UHFFFAOYSA-K phosphate Chemical compound [O-]P([O-])([O-])=O NBIIXXVUZAFLBC-UHFFFAOYSA-K 0.000 description 1
- 229940085991 phosphate ion Drugs 0.000 description 1
- 230000035484 reaction time Effects 0.000 description 1
- 125000005920 sec-butoxy group Chemical group 0.000 description 1
- 230000001235 sensitizing effect Effects 0.000 description 1
- 238000000926 separation method Methods 0.000 description 1
- 125000001174 sulfone group Chemical group 0.000 description 1
- 238000001308 synthesis method Methods 0.000 description 1
- 238000003786 synthesis reaction Methods 0.000 description 1
Landscapes
- Pyridine Compounds (AREA)
- Low-Molecular Organic Synthesis Reactions Using Catalysts (AREA)
Abstract
Description
本発明は複素環化合物の製造方法、特にモノハロゲノ置換ピリジン化合物の製造方法に関する。 The present invention relates to a method for producing a heterocyclic compound, and more particularly to a method for producing a monohalogeno-substituted pyridine compound.
異なる置換基を有する2,6−二置換ピリジン化合物は、耐光堅牢度に優れた繊維用分散染料や、写真用転化剤、増感色素、電子材料、医農薬等に使用する化合物またはその合成中間体として幅広く用いられている。 2,6-disubstituted pyridine compounds having different substituents are disperse dyes for fibers excellent in light fastness, compounds used for photographic conversion agents, sensitizing dyes, electronic materials, medical pesticides, etc. Widely used as a body.
従来の製造方法として、例えば、特許文献1および特許文献2には2,6−ジクロロ−3−シアノ−4−メチルピリジンから染料を合成する方法等が記載されているが、特許文献1の方法では溶剤を使用しており、特許文献2では試薬を過剰に使用することにより溶剤を兼ねて合成を行っており、コストが高くなる方法であり、また、作業安全上からも問題の多い方法であった。 As a conventional production method, for example, Patent Document 1 and Patent Document 2 describe a method of synthesizing a dye from 2,6-dichloro-3-cyano-4-methylpyridine. In US Pat. No. 6,099,097, a solvent is used, and in Patent Document 2, synthesis is performed also as a solvent by using an excessive amount of reagent, which is a costly method and a method with many problems from the viewpoint of work safety. there were.
高収率、高純度且つ安価に2,6−二置換ピリジン化合物を製造するためのモノハロゲノ置換ピリジン化合物の製造方法であって、引火点の低い危険物や燃焼性のある薬品等の使用を極力抑え、水中の反応系で実施する製造方法を提供すること。 A method for producing a monohalogeno-substituted pyridine compound for producing a 2,6-disubstituted pyridine compound at a high yield, high purity and low cost, and to use as much as possible a dangerous substance having a low flash point or a flammable chemical. To provide a production method that is carried out in an underwater reaction system.
本発明者らは前記課題を解決すべく鋭意研究を重ねた結果、特に2位と6位にハロゲン原子を有するピリジン化合物とアニリン誘導体とを、水中で置換反応させることで、高収率、高純度、安価且つ安全に、モノハロゲノ置換ピリジン化合物を製造出来ることを見出し、本発明を完成させた。 As a result of intensive studies to solve the above problems, the present inventors have carried out a substitution reaction in water with a pyridine compound having a halogen atom at the 2nd and 6th positions, and an aniline derivative. The present inventors have found that a monohalogeno-substituted pyridine compound can be produced with purity, low cost and safety, and completed the present invention.
即ち、本発明は以下の1)〜4)に関する。
1)下記式(1)で表される脱離基として2個のハロゲン原子を有する複素環化合物に下記式(2)で表されるアニリン誘導体を水中で置換反応させることを特徴とする下記式(3)で表されるモノハロゲノ置換複素環化合物の製造方法。
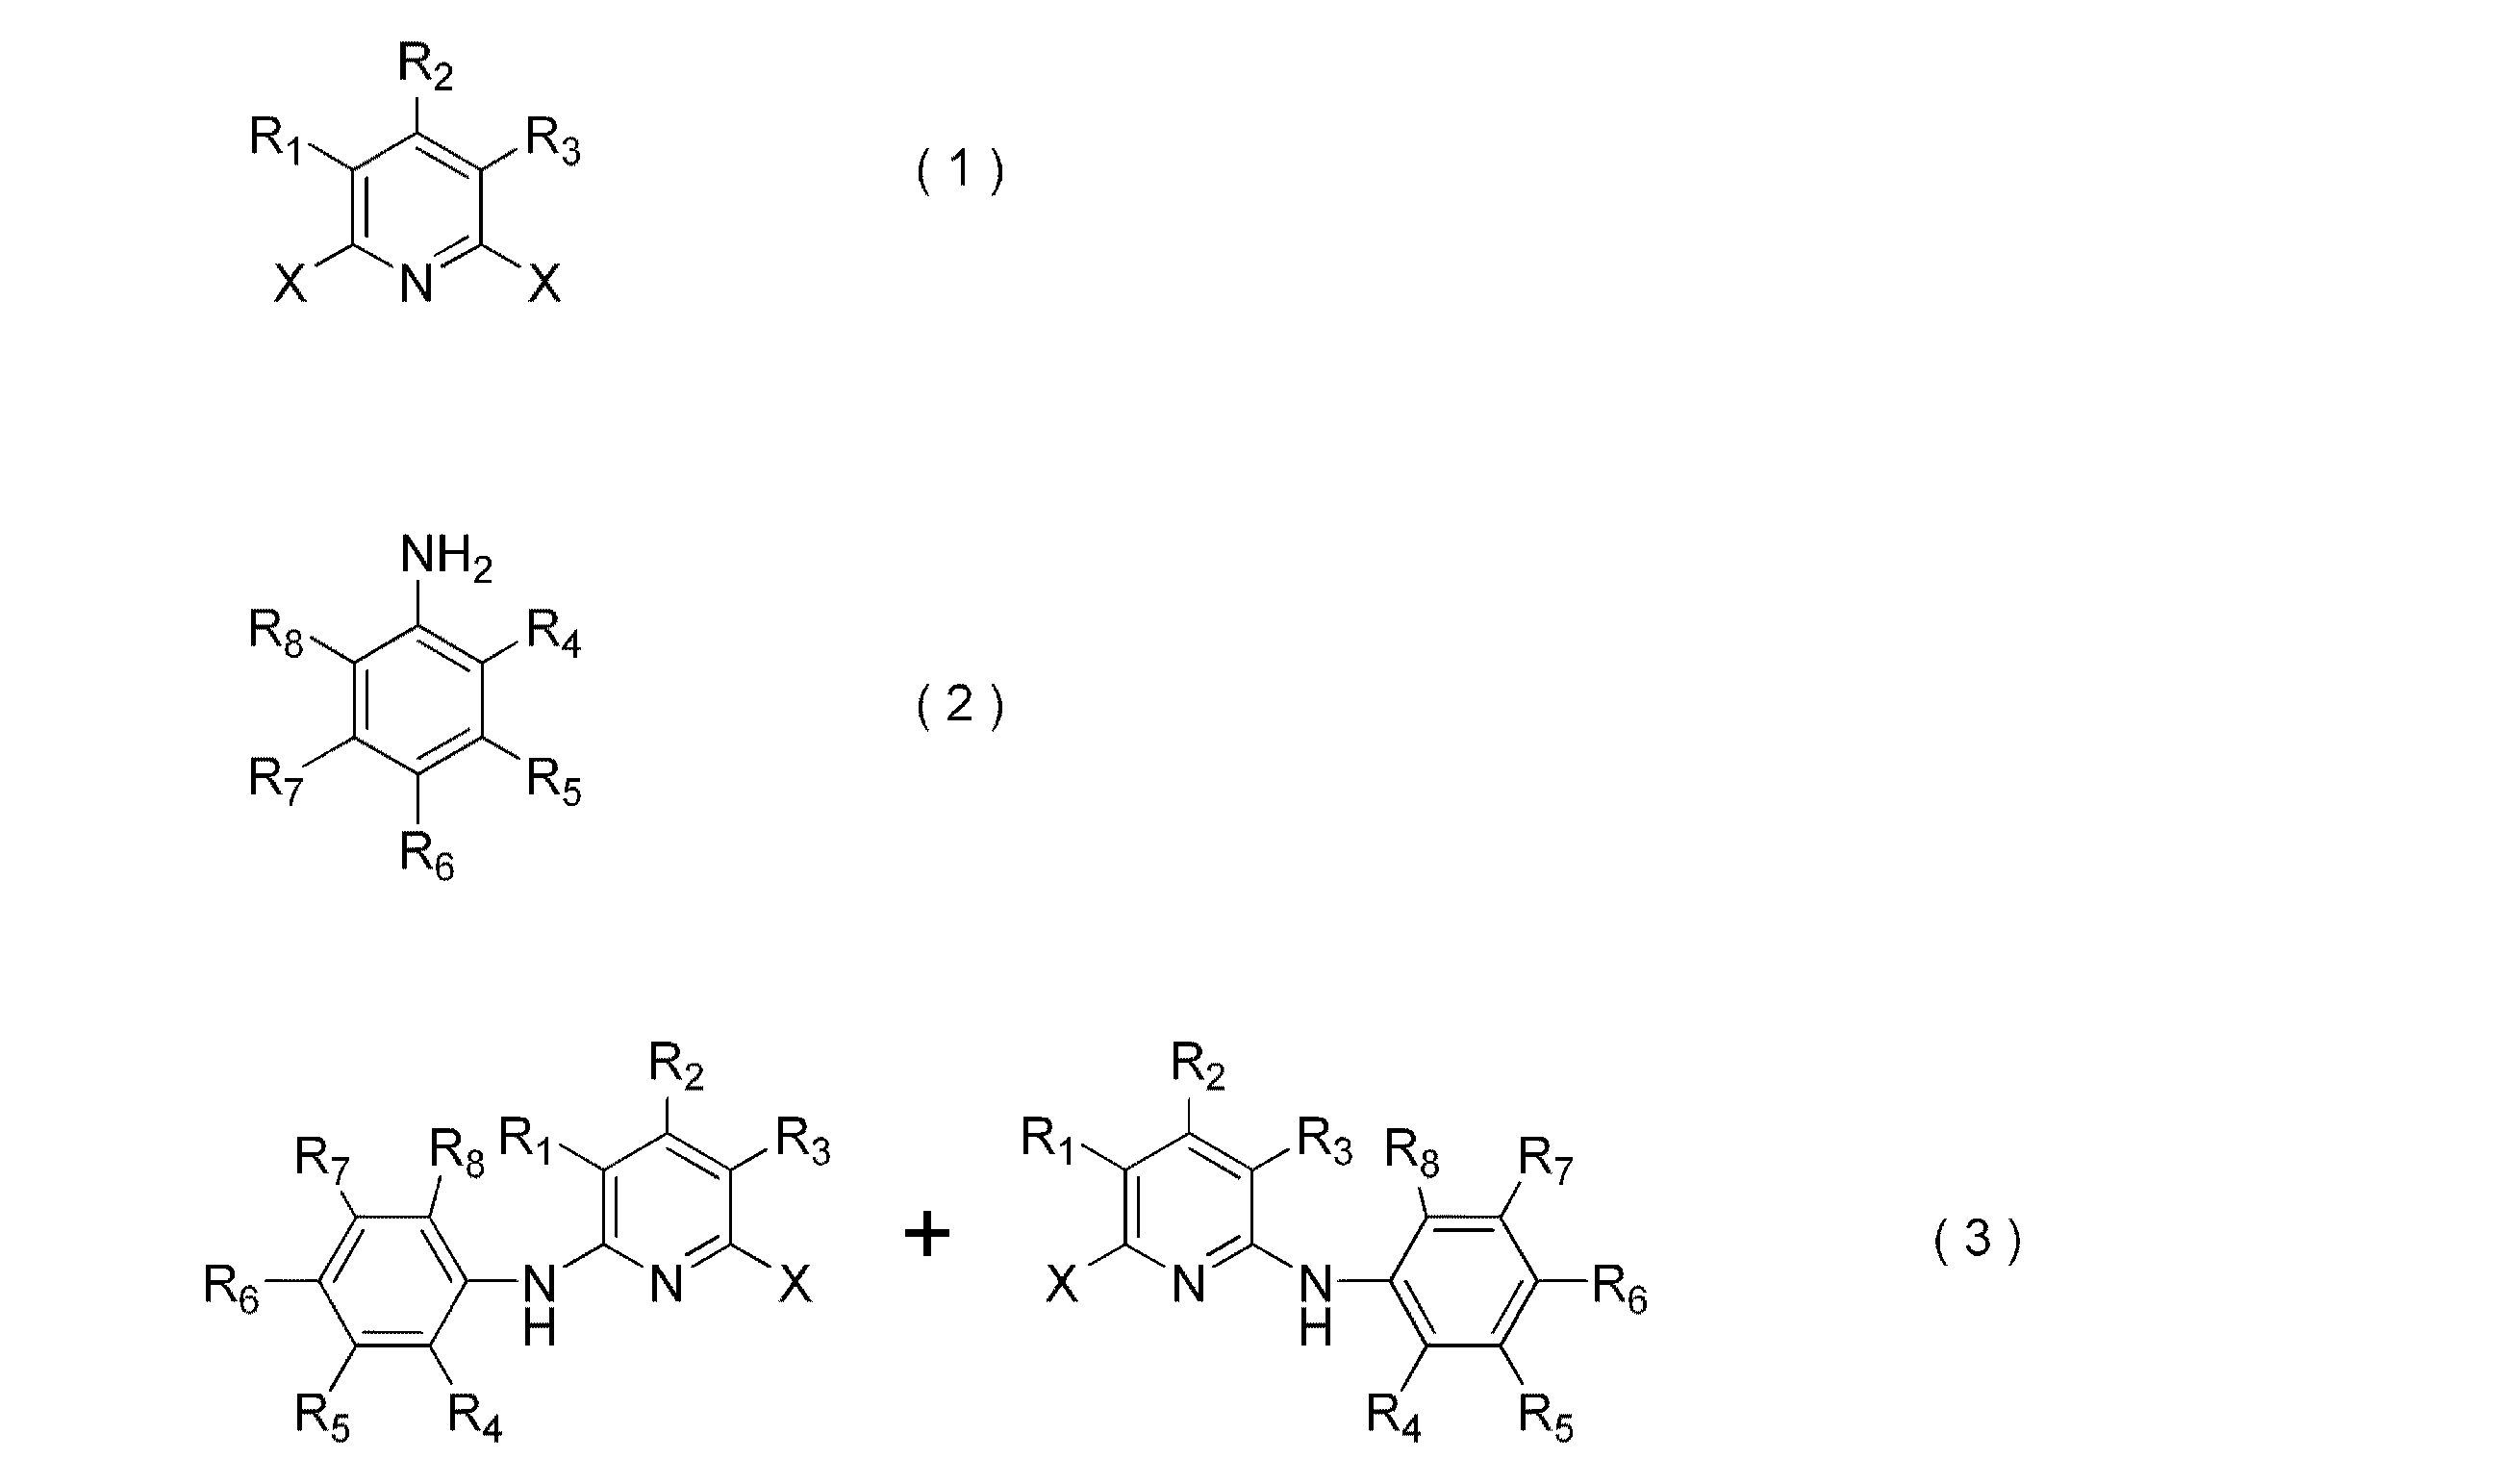
1) Substitution reaction of an aniline derivative represented by the following formula (2) with a heterocyclic compound having two halogen atoms as a leaving group represented by the following formula (1) in water The manufacturing method of the monohalogeno substituted heterocyclic compound represented by (3).
2)Xが塩素原子、R1がシアノ基、R2が(C1〜C3)アルキル基、R3およびR4〜R8が水素原子である前記1)記載の製造方法。
3)置換反応に、炭酸カリウム、炭酸ナトリウム、炭酸水素ナトリウムおよび炭酸カルシウムからなる無機塩基群から選ばれる少なくとも一つの塩基を使用することを特徴とする前記1)または2)に記載の製造方法。
4)置換反応の反応促進用触媒として、酸化亜鉛、ヨウ化銅および水酸化ナトリウムからなる群から選ばれる少なくとも一つの化合物を使用することを特徴とする前記3)記載の製造方法。
2) The production method according to 1), wherein X is a chlorine atom, R 1 is a cyano group, R 2 is a (C1-C3) alkyl group, and R 3 and R 4 to R 8 are hydrogen atoms.
3) The production method according to 1) or 2) above, wherein at least one base selected from an inorganic base group consisting of potassium carbonate, sodium carbonate, sodium hydrogen carbonate and calcium carbonate is used for the substitution reaction.
4) The production method according to 3) above, wherein at least one compound selected from the group consisting of zinc oxide, copper iodide and sodium hydroxide is used as a catalyst for promoting the reaction of the substitution reaction.
本発明の水を溶媒として使用することを特徴とする製造方法により高収率、高純度、安価且つ安全にモノハロゲノ置換ピリジン化合物を製造することが可能となった。 The production method characterized by using the water of the present invention as a solvent makes it possible to produce a monohalogeno-substituted pyridine compound with high yield, high purity, low cost and safety.
以下に、本発明を詳細に説明する。
本発明の前記式(3)で表されるモノハロゲノ置換複素環化合物の製造方法は、前記式(1)で表される脱離基として2個のハロゲン原子を有する複素環化合物に前記式(2)で表されるアニリン誘導体を水中で置換反応させることを特徴とする[式中、Xはそれぞれ独立に塩素原子、臭素原子またはヨウ素原子を表し、R1〜R3はそれぞれ独立に水素原子、(C1〜C6)アルキル基、フェニル基、シアノ基またはニトロ基を表し、R4〜R8はそれぞれ独立に水素原子、(C1〜C6)アルキル基、(C1〜C6)アルコキシ基、(C1〜C6)アルコキシ(C1〜C6)アルコキシ基、置換基を有していてもよいフェニル基を表す。]
The present invention is described in detail below.
In the method for producing a monohalogeno-substituted heterocyclic compound represented by the formula (3) of the present invention, the heterocyclic compound having two halogen atoms as the leaving group represented by the formula (1) is added to the formula (2). The aniline derivative represented by the above formula is substituted in water [wherein, X represents independently a chlorine atom, a bromine atom or an iodine atom, R 1 to R 3 each independently represents a hydrogen atom, (C1-C6) represents an alkyl group, a phenyl group, a cyano group, or a nitro group, and R 4 to R 8 each independently represent a hydrogen atom, (C1-C6) alkyl group, (C1-C6) alkoxy group, (C1-C6) C6) represents an alkoxy (C1-C6) alkoxy group and a phenyl group which may have a substituent. ]
。
まず、本発明の製造方法で製造される前記式(3)で表されるモノハロゲノ置換複素環化合物、即ち、モノハロゲノ置換ピリジン化合物について説明する。
.
First, the monohalogeno-substituted heterocyclic compound represented by the formula (3) produced by the production method of the present invention, that is, the monohalogeno-substituted pyridine compound will be described.
R1〜R3における(C1〜C6)アルキル基としては、直鎖、分岐のアルキル基が挙げられ、例えば、メチル基、エチル基、n−プロピル基、イソプロピル基、n−ブチル基、t−ブチル基、n−ヘプチル基、n−ヘキシル基等が挙げられる。
好ましくはR1がシアノ基、R2が(C1〜C3)アルキル基、例えば、メチル基、エチル基、n−プロピル基、イソプロピル基、R3が水素原子であり、中でも特にR2がメチル基である化合物が好ましい。
Examples of the (C1 to C6) alkyl group in R 1 to R 3 include linear and branched alkyl groups. For example, a methyl group, an ethyl group, an n-propyl group, an isopropyl group, an n-butyl group, t- A butyl group, n-heptyl group, n-hexyl group, etc. are mentioned.
Preferably, R 1 is a cyano group, R 2 is a (C1-C3) alkyl group, for example, a methyl group, an ethyl group, an n-propyl group, an isopropyl group, R 3 is a hydrogen atom, and in particular, R 2 is a methyl group. Is preferred.
R4〜R8における(C1〜C6)アルキル基としては、R1〜R3における(C1〜C6)アルキル基と同様な基が挙げられる。(C1〜C6)アルコキシ基としては、例えば、メトキシ基、エトキシ基、n−プロポキシ基、iso−プロポキシ基、n−ブトキシ基、iso−ブトキシ基、sec−ブトキシ基、n−ペントキシ基、メチルブトキシ基、エチルプロポキシ基、エチルブトキシ基等が挙げられる。(C1〜C6)アルコキシ(C1〜C6)アルコキシ基としてはメトキシメトキシ基、メトキシエトキシ基、メトキシプロポキシ基、エトキシメトキシ基、エトキシエトキシ基、エトキシプロポキシ基等が挙げられる。置換基を有していてもよいフェニル基の置換基としては、ハロゲン原子、ヒドロキシ基、カルボキシ基、スルホン基、フェニル基等が挙げられる。
好ましくはR4〜R8が水素原子の化合物である。
Examples of the (C1 to C6) alkyl group for R 4 to R 8 include the same groups as the (C1 to C6) alkyl group for R 1 to R 3 . Examples of the (C1 to C6) alkoxy group include a methoxy group, an ethoxy group, an n-propoxy group, an iso-propoxy group, an n-butoxy group, an iso-butoxy group, a sec-butoxy group, an n-pentoxy group, and a methylbutoxy group. Group, ethylpropoxy group, ethylbutoxy group and the like. (C1-C6) alkoxy (C1-C6) alkoxy groups include methoxymethoxy, methoxyethoxy, methoxypropoxy, ethoxymethoxy, ethoxyethoxy, ethoxypropoxy and the like. Examples of the substituent of the phenyl group which may have a substituent include a halogen atom, a hydroxy group, a carboxy group, a sulfone group, and a phenyl group.
R 4 to R 8 are preferably hydrogen atom compounds.
式(3)で表される化合物は、その置換基の種類に起因する互変異性体や位置異性体が存在する場合があるが、本発明にはそれらの異性体およびそれら異性体の混合物はすべて包含される。 In the compound represented by the formula (3), there may be a tautomer or a positional isomer due to the type of the substituent. In the present invention, these isomers and mixtures of these isomers are All are included.
本発明において前記式(3)で表されるモノハロゲノ置換複素環化合物は、塩を形成していてもよく、該塩も本発明に含まれる。該塩は特に限定されず、例えば、ハロゲン化物イオン、硫酸イオン、炭酸イオン、硝酸イオン、スルホン酸イオン、リン酸イオン、酢酸イオン等の陰イオンとの塩、金属イオン、アンモニウムイオン等の陽イオンとの塩が挙げられる。また、置換基によっては分子内塩を形成していてもよい。 In the present invention, the monohalogeno-substituted heterocyclic compound represented by the formula (3) may form a salt, and the salt is also included in the present invention. The salt is not particularly limited, for example, a salt with an anion such as halide ion, sulfate ion, carbonate ion, nitrate ion, sulfonate ion, phosphate ion, acetate ion, cation such as metal ion, ammonium ion, etc. And the salt. Further, depending on the substituent, an inner salt may be formed.
次に、本発明の製造方法について詳しく述べる。
本発明の製造方法に用いられる式(2)で表されるアニリン誘導体におけるR4〜R8は前記の式(3)で表されるモノハロゲノ置換ピリジン化合物におけるR4〜R8と同様であり、好ましい基も同様である。その使用量は適宜選択可能であるが、通常は式(1)で表される複素環化合物1モルに対して0.5〜30モル程度用いることが出来、より好ましくは1〜10モル程度であり、特に好ましくは1.5〜5モル程度である。
Next, the production method of the present invention will be described in detail.
R 4 to R 8 in the aniline derivative represented by the formula (2) used in the production method of the present invention are the same as R 4 to R 8 in the monohalogeno-substituted pyridine compound represented by the formula (3), The preferable group is also the same. The amount used can be selected as appropriate, but usually it can be used in an amount of about 0.5 to 30 mol, more preferably about 1 to 10 mol, per 1 mol of the heterocyclic compound represented by the formula (1). Yes, particularly preferably about 1.5 to 5 mol.
本発明の製造方法に用いられる式(1)で表される複素環化合物におけるXは塩素原子、臭素原子、ヨウ素原子が挙げられ、中でも好ましくは2個のXがともに塩素原子である化合物が挙げられる。R1〜R3は前記の式(3)で表されるモノハロゲノ置換複素環化合物におけるR1〜R3と同様であり、好ましい基も同様である。
反応溶媒としては水のみを使用することが好ましく、その使用量は特に限定されず、反応原料に応じて適宜選択することが出来るが、通常は式(1)で表される複素環化合物に対して質量比で0.1〜100倍程度、好ましくは0.5〜10倍程度、特に好ましくは1〜5倍程度である。
Examples of X in the heterocyclic compound represented by the formula (1) used in the production method of the present invention include a chlorine atom, a bromine atom and an iodine atom. Among them, a compound in which two Xs are both chlorine atoms is preferable. It is done. R 1 to R 3 is the same as R 1 to R 3 in the monohalogenoacetic substituted heterocyclic compound represented by the formula (3), is the same preferable groups.
As the reaction solvent, it is preferable to use only water, and the amount used is not particularly limited and can be appropriately selected according to the reaction raw material, but is usually based on the heterocyclic compound represented by the formula (1). The mass ratio is about 0.1 to 100 times, preferably about 0.5 to 10 times, and particularly preferably about 1 to 5 times.
本発明の製造方法における反応温度は特に限定されず、適宜選択することが出来るが、通常は0〜150℃程度であり、好ましくは40〜120℃程度、特に好ましくは80〜105℃程度である。 The reaction temperature in the production method of the present invention is not particularly limited and can be appropriately selected, but is usually about 0 to 150 ° C, preferably about 40 to 120 ° C, particularly preferably about 80 to 105 ° C. .
本発明の製造方法における反応時間は特に限定されず、反応原料に応じて適宜選択することが出来るが、通常は1分〜50時間程度であり、好ましくは5〜40時間程度、特に好ましくは10〜30時間程度である。
HPLC(高分解能液体クロマログラフィー)にて原料の消失により反応の終了を確認することも出来る。
The reaction time in the production method of the present invention is not particularly limited and can be appropriately selected depending on the reaction raw materials. Usually, it is about 1 minute to 50 hours, preferably about 5 to 40 hours, particularly preferably 10 ~ 30 hours.
Completion of the reaction can be confirmed by disappearance of the raw material by HPLC (high resolution liquid chromatography).
本発明の製造方法では炭酸カリウム、炭酸ナトリウム、炭酸水素ナトリウムおよび炭酸カルシウムからなる無機塩基群から選ばれる少なくとも一つの塩基を使用するのが好ましい。無機塩基を使用する場合、その使用量は前記式(1)で表される複素環化合物1モルに対して0.1〜30モル程度、より好ましくは0.5〜10モル程度、特に好ましくは1〜3モル程度である。
本発明の製造方法ではさらに反応促進用触媒として、酸化亜鉛、ヨウ化銅および水酸化ナトリウムからなる群から選ばれる少なくとも一つの化合物を使用するのが好ましい。反応促進用触媒を使用する場合、その使用量は前記式(1)で表される複素環化合物1モルに対して0.01〜10モル程度、より好ましくは0.1〜5モル程度、特に好ましくは0.2〜1モル程度である。
In the production method of the present invention, it is preferable to use at least one base selected from the group of inorganic bases consisting of potassium carbonate, sodium carbonate, sodium hydrogen carbonate and calcium carbonate. When an inorganic base is used, the amount used is about 0.1 to 30 mol, more preferably about 0.5 to 10 mol, particularly preferably about 1 mol of the heterocyclic compound represented by the formula (1). About 1 to 3 moles.
In the production method of the present invention, it is preferable to use at least one compound selected from the group consisting of zinc oxide, copper iodide and sodium hydroxide as the reaction promoting catalyst. When using the reaction promoting catalyst, the amount used is about 0.01 to 10 mol, more preferably about 0.1 to 5 mol, particularly about 1 to 5 mol, based on 1 mol of the heterocyclic compound represented by the formula (1). Preferably it is about 0.2-1 mol.
式(1)で表される複素環化合物や式(2)で表されるアニリン誘導体、さらに、無機塩基、反応促進用触媒および反応溶媒の反応系内への添加順序は任意であってよく特に限定されない。 The order of addition of the heterocyclic compound represented by the formula (1), the aniline derivative represented by the formula (2), the inorganic base, the reaction promoting catalyst and the reaction solvent into the reaction system may be arbitrary. It is not limited.
以下、実施例により本発明をさらに詳細に説明する。実施例において、特に断りのない限り、部は重量部を、%は重量%を意味する。
実施例1
2,6−ジクロロ−4−メチル−3−ピリジンカルボニトリル(前記式(1)でXが塩素原子、R1がシアノ基、R2がメチル基、R3が水素原子;30g、0.16mol)、アニリン(前記式(2)でR4〜R8が水素原子;45g、0.48mol)、水(30g)を混合し、加熱昇温して95〜105℃で15時間加熱攪拌した。これを冷却し、得られた結晶をろ取し、水でかけ洗いした。得られた結晶に対し、攪拌しながら10%塩酸水溶液を滴下し、1時間攪拌した後、得られた結晶をろ取し、3%塩酸水でかけ洗い、続いて水でろ液が中性になるまでかけ洗いした。これを乾燥して主生成物1、主生成物2、副生成物の混合物(37.7g、96%)を得た。
Example 1
2,6-dichloro-4-methyl-3-pyridinecarbonitrile (in the above formula (1), X is a chlorine atom, R 1 is a cyano group, R 2 is a methyl group, R 3 is a hydrogen atom; 30 g, 0.16 mol) ) And aniline (in formula (2), R 4 to R 8 are hydrogen atoms; 45 g, 0.48 mol) and water (30 g) were mixed, heated to an elevated temperature, and stirred at 95 to 105 ° C. for 15 hours. This was cooled, and the resulting crystals were collected by filtration and washed with water. A 10% aqueous hydrochloric acid solution is added dropwise to the obtained crystals while stirring, and after stirring for 1 hour, the obtained crystals are collected by filtration, washed with 3% aqueous hydrochloric acid, and the filtrate becomes neutral with water. Until it was washed. This was dried to obtain a mixture of main product 1, main product 2 and by-products (37.7 g, 96%).
実施例2
2,6−ジクロロ−4−メチル−3−ピリジンカルボニトリル(30g、0.16mol)、アニリン(22g、0.24mol)、炭酸カリウム(33g、0.24mol)、水(60g)を混合し、加熱昇温して98〜110℃で5時間加熱攪拌後、48%水酸化ナトリウム水溶液(6.7g)を添加し、さらに10時間加熱攪拌した。これを冷却し、得られた結晶をろ取し、水でかけ洗いした。得られた結晶に対し、攪拌しながら10%塩酸水溶液を滴下し、1時間攪拌した後、得られた結晶をろ取し、3%塩酸水でかけ洗い、続いて水でろ液が中性になるまでかけ洗いした。これを乾燥して主生成物1、主生成物2、副生成物の混合物(36.7g、94%)を得た。
Example 2
2,6-dichloro-4-methyl-3-pyridinecarbonitrile (30 g, 0.16 mol), aniline (22 g, 0.24 mol), potassium carbonate (33 g, 0.24 mol), water (60 g) were mixed, The mixture was heated and heated and stirred at 98 to 110 ° C. for 5 hours, 48% aqueous sodium hydroxide solution (6.7 g) was added, and the mixture was further stirred for 10 hours. This was cooled, and the resulting crystals were collected by filtration and washed with water. A 10% aqueous hydrochloric acid solution is added dropwise to the obtained crystals while stirring, and after stirring for 1 hour, the obtained crystals are collected by filtration, washed with 3% aqueous hydrochloric acid, and the filtrate becomes neutral with water. Until it was washed. This was dried to obtain a mixture of main product 1, main product 2, and by-products (36.7 g, 94%).
実施例3
2,6−ジクロロ−4−メチル−3−ピリジンカルボニトリル(20g、0.11mol)、アニリン(20g、0.21mol)、炭酸カリウム(22g、0.16mol)、水(40g)、酸化亜鉛(0.5g、0.006mol)を混合し、加熱昇温して95〜105℃で15時間加熱攪拌した。これを冷却し、得られた結晶をろ取し、水でかけ洗いした。得られた結晶に対し、攪拌しながら10%塩酸水溶液を滴下し、1時間攪拌した後、得られた結晶をろ取し、3%塩酸水でかけ洗い、続いて水でろ液が中性になるまでかけ洗いした。これを乾燥して主生成物1、主生成物2、副生成物の混合物(25.4g、97%)を得た。
Example 3
2,6-dichloro-4-methyl-3-pyridinecarbonitrile (20 g, 0.11 mol), aniline (20 g, 0.21 mol), potassium carbonate (22 g, 0.16 mol), water (40 g), zinc oxide ( 0.5 g, 0.006 mol) was mixed, heated and heated, and stirred at 95 to 105 ° C. for 15 hours. This was cooled, and the resulting crystals were collected by filtration and washed with water. A 10% aqueous hydrochloric acid solution is added dropwise to the obtained crystals while stirring, and after stirring for 1 hour, the obtained crystals are collected by filtration, washed with 3% aqueous hydrochloric acid, and the filtrate becomes neutral with water. Until it was washed. This was dried to obtain a mixture of main product 1, main product 2 and by-products (25.4 g, 97%).
比較例1
2,6−ジクロロ−4−メチル−3−ピリジンカルボニトリル(50g、0.27mol)、アニリン(80.4g、0.86mol)、イソプロパノール(54g)を混合し、加熱昇温して80〜95℃で10時間加熱攪拌した。これを冷却し、冷やした30%塩酸水溶液に注いで晶析した。これをろ取し、水でろ液が中性になるまでかけ洗いした。これを乾燥して主生成物1、主生成物2、副生成物の混合物(63.6g、98%)を得た。
Comparative Example 1
2,6-dichloro-4-methyl-3-pyridinecarbonitrile (50 g, 0.27 mol), aniline (80.4 g, 0.86 mol) and isopropanol (54 g) were mixed, heated to 80-95 by heating. The mixture was stirred at 10 ° C. for 10 hours. This was cooled and poured into a cooled 30% aqueous hydrochloric acid solution for crystallization. This was collected by filtration and washed with water until the filtrate became neutral. This was dried to obtain a mixture of main product 1, main product 2 and by-products (63.6 g, 98%).
次に、得られた混合物のHPLC分析を行い、主生成物1、主生成物2、副生成物の組成比を決定した。 Next, HPLC analysis of the obtained mixture was performed, and the composition ratio of the main product 1, the main product 2, and the by-product was determined.
表1
HPLC分析条件
モデル:島津 Prominence
カラム:Inertsil ODS−3(5μm) φ4.6x150mm (40℃)
検出器:UV210nm
移動相および分離条件:
A:アセトニトリル B:水
A液30%→(15分)→100%→(5分)
HPLC analysis condition model: Shimazu Prominence
Column: Inertsil ODS-3 (5 μm) φ4.6 × 150 mm (40 ° C.)
Detector: UV210nm
Mobile phase and separation conditions:
A: Acetonitrile B: Water A solution 30% → (15 minutes) → 100% → (5 minutes)
表1から明らかなように有機溶媒を使用しない本発明の製造方法によれば副生成物の割合が5%以下と少なく、目的とする化合物の混合物が高収率、高純度で得られる。
先にも述べたが、従来の製造法として知られている特許文献1に記載の溶媒にイソプロパノールを用いて合成する方法(比較例1)では副生成物の割合が11%であり、生成物は目的とする化合物の混合物の純度が低い。また、特許文献2に記載の試薬を大過剰量使用して合成する方法でも望まない副生成物が多く生成した。
As is apparent from Table 1, according to the production method of the present invention in which no organic solvent is used, the ratio of by-products is as low as 5% or less, and a desired mixture of compounds can be obtained in high yield and high purity.
As described above, in the method of synthesizing using isopropanol in the solvent described in Patent Document 1 known as a conventional production method (Comparative Example 1), the by-product ratio is 11%, and the product The purity of the target compound mixture is low. In addition, a large amount of undesired by-products were produced even in the synthesis method using a large excess of the reagent described in Patent Document 2.
Claims (4)
Priority Applications (1)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| JP2011117605A JP5748210B2 (en) | 2011-05-26 | 2011-05-26 | Method for producing heterocyclic compound |
Applications Claiming Priority (1)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| JP2011117605A JP5748210B2 (en) | 2011-05-26 | 2011-05-26 | Method for producing heterocyclic compound |
Publications (2)
| Publication Number | Publication Date |
|---|---|
| JP2012246231A true JP2012246231A (en) | 2012-12-13 |
| JP5748210B2 JP5748210B2 (en) | 2015-07-15 |
Family
ID=47467056
Family Applications (1)
| Application Number | Title | Priority Date | Filing Date |
|---|---|---|---|
| JP2011117605A Expired - Fee Related JP5748210B2 (en) | 2011-05-26 | 2011-05-26 | Method for producing heterocyclic compound |
Country Status (1)
| Country | Link |
|---|---|
| JP (1) | JP5748210B2 (en) |
Cited By (1)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| CN119528808A (en) * | 2024-11-25 | 2025-02-28 | 浙江闰土股份有限公司 | High-sun-durability red dye intermediate and preparation method thereof |
Citations (7)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| JPS4962477A (en) * | 1972-06-22 | 1974-06-17 | ||
| JPS4994677A (en) * | 1972-12-16 | 1974-09-09 | ||
| JPS6139347B2 (en) * | 1972-03-10 | 1986-09-03 | Basf Ag | |
| JPH08507496A (en) * | 1991-12-20 | 1996-08-13 | ビーエーエスエフ アクチェンゲゼルシャフト | Diaminopyridine manufacturing method |
| JP2006265207A (en) * | 2005-03-25 | 2006-10-05 | Fuji Photo Film Co Ltd | Method for producing heterocyclic compound |
| JP2009524670A (en) * | 2006-01-27 | 2009-07-02 | アレイ バイオファーマ、インコーポレイテッド | Glucokinase activator |
| JP2010529178A (en) * | 2007-06-13 | 2010-08-26 | サノフイ−アベンテイス | Derivatives of 7-alkynyl-1,8-naphthyridone, methods for their preparation and their use in therapy |
-
2011
- 2011-05-26 JP JP2011117605A patent/JP5748210B2/en not_active Expired - Fee Related
Patent Citations (7)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| JPS6139347B2 (en) * | 1972-03-10 | 1986-09-03 | Basf Ag | |
| JPS4962477A (en) * | 1972-06-22 | 1974-06-17 | ||
| JPS4994677A (en) * | 1972-12-16 | 1974-09-09 | ||
| JPH08507496A (en) * | 1991-12-20 | 1996-08-13 | ビーエーエスエフ アクチェンゲゼルシャフト | Diaminopyridine manufacturing method |
| JP2006265207A (en) * | 2005-03-25 | 2006-10-05 | Fuji Photo Film Co Ltd | Method for producing heterocyclic compound |
| JP2009524670A (en) * | 2006-01-27 | 2009-07-02 | アレイ バイオファーマ、インコーポレイテッド | Glucokinase activator |
| JP2010529178A (en) * | 2007-06-13 | 2010-08-26 | サノフイ−アベンテイス | Derivatives of 7-alkynyl-1,8-naphthyridone, methods for their preparation and their use in therapy |
Cited By (1)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| CN119528808A (en) * | 2024-11-25 | 2025-02-28 | 浙江闰土股份有限公司 | High-sun-durability red dye intermediate and preparation method thereof |
Also Published As
| Publication number | Publication date |
|---|---|
| JP5748210B2 (en) | 2015-07-15 |
Similar Documents
| Publication | Publication Date | Title |
|---|---|---|
| EP3882236B1 (en) | Production method for 2,5-dicyano-3,6-dihalogenopyrazine | |
| AU2014373959A1 (en) | 5-fluoro-4-imino-3-(alkyl/substituted alkyl)-1- (arylsulfonyl)-3,4-dihydropyrimidin-2(1H)-one and processes for their preparation | |
| EP3452437B1 (en) | Method for aromatic fluorination | |
| CN109053505B (en) | A kind of synthetic method of important intermediate of filocoxib | |
| US12071399B2 (en) | Process for the preparation of arylsulfonylpropenenitriles | |
| JP5748210B2 (en) | Method for producing heterocyclic compound | |
| JP2005536558A (en) | Process for the production of nitrooxy derivatives of naproxen | |
| RU2635659C1 (en) | Method of producing benzil ether derivatives of 2-aminonicotinic acid | |
| KR20120071339A (en) | Process for the production of bendamustine alkyl ester, bendamustine, and derivatives thereof | |
| WO2019141230A1 (en) | Processes for the synthesis of sulfentrazone | |
| CN102260287B (en) | Preparation method of herbicide anilofos | |
| EP1985610A1 (en) | Improved method for producing nitroisourea derivative | |
| JP6166996B2 (en) | Method for producing hydrazinoarylcarboxylic acids | |
| JP2003335735A (en) | Method for producing perfluoroisopropylaniline | |
| CN106749017A (en) | A kind of synthetic method of 1,8 naphthalimides of 3,6 halogen atoms substitution | |
| EP2819983A1 (en) | Process for the preparation of substituted phenylpropanones | |
| DE102009045366A1 (en) | Polycyclic pentafluorosulfanylbenzene compound and process for the preparation of the compound | |
| JP4138067B2 (en) | Method for producing methine derivative | |
| JP6477187B2 (en) | Process for producing 2-amino-6-methylnicotinic acid ester | |
| KR20200095741A (en) | Process for the preparation of acyloxybenzenesulfonates | |
| JP3646224B2 (en) | Method for producing benzoylacetonitrile derivative | |
| EP4372056A1 (en) | Xanthene compound and method for producing same | |
| JP7035815B2 (en) | Method for Producing Trivalent Hypervalent Iodine Compound Using Hypochlorite | |
| JP2018162218A (en) | Novel cyclic urea derivatives-hydrobromide | |
| RU2702121C1 (en) | Method of producing 2-amino-nicotinic acid benzyl ester derivative |
Legal Events
| Date | Code | Title | Description |
|---|---|---|---|
| A621 | Written request for application examination |
Free format text: JAPANESE INTERMEDIATE CODE: A621 Effective date: 20140127 |
|
| A977 | Report on retrieval |
Free format text: JAPANESE INTERMEDIATE CODE: A971007 Effective date: 20141023 |
|
| A131 | Notification of reasons for refusal |
Free format text: JAPANESE INTERMEDIATE CODE: A131 Effective date: 20141110 |
|
| A521 | Written amendment |
Free format text: JAPANESE INTERMEDIATE CODE: A523 Effective date: 20150108 |
|
| TRDD | Decision of grant or rejection written | ||
| A01 | Written decision to grant a patent or to grant a registration (utility model) |
Free format text: JAPANESE INTERMEDIATE CODE: A01 Effective date: 20150507 |
|
| A61 | First payment of annual fees (during grant procedure) |
Free format text: JAPANESE INTERMEDIATE CODE: A61 Effective date: 20150507 |
|
| R150 | Certificate of patent or registration of utility model |
Ref document number: 5748210 Country of ref document: JP Free format text: JAPANESE INTERMEDIATE CODE: R150 |
|
| LAPS | Cancellation because of no payment of annual fees |