JP7502425B2 - BRM TARGETING COMPOUNDS AND RELATED METHODS OF USE - Google Patents
BRM TARGETING COMPOUNDS AND RELATED METHODS OF USE Download PDFInfo
- Publication number
- JP7502425B2 JP7502425B2 JP2022520255A JP2022520255A JP7502425B2 JP 7502425 B2 JP7502425 B2 JP 7502425B2 JP 2022520255 A JP2022520255 A JP 2022520255A JP 2022520255 A JP2022520255 A JP 2022520255A JP 7502425 B2 JP7502425 B2 JP 7502425B2
- Authority
- JP
- Japan
- Prior art keywords
- optionally substituted
- ulm
- alkyl
- group
- groups
- Prior art date
- Legal status (The legal status is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the status listed.)
- Active
Links
Classifications
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D417/00—Heterocyclic compounds containing two or more hetero rings, at least one ring having nitrogen and sulfur atoms as the only ring hetero atoms, not provided for by group C07D415/00
- C07D417/14—Heterocyclic compounds containing two or more hetero rings, at least one ring having nitrogen and sulfur atoms as the only ring hetero atoms, not provided for by group C07D415/00 containing three or more hetero rings
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61K—PREPARATIONS FOR MEDICAL, DENTAL OR TOILETRY PURPOSES
- A61K47/00—Medicinal preparations characterised by the non-active ingredients used, e.g. carriers or inert additives; Targeting or modifying agents chemically bound to the active ingredient
- A61K47/50—Medicinal preparations characterised by the non-active ingredients used, e.g. carriers or inert additives; Targeting or modifying agents chemically bound to the active ingredient the non-active ingredient being chemically bound to the active ingredient, e.g. polymer-drug conjugates
- A61K47/51—Medicinal preparations characterised by the non-active ingredients used, e.g. carriers or inert additives; Targeting or modifying agents chemically bound to the active ingredient the non-active ingredient being chemically bound to the active ingredient, e.g. polymer-drug conjugates the non-active ingredient being a modifying agent
- A61K47/54—Medicinal preparations characterised by the non-active ingredients used, e.g. carriers or inert additives; Targeting or modifying agents chemically bound to the active ingredient the non-active ingredient being chemically bound to the active ingredient, e.g. polymer-drug conjugates the non-active ingredient being a modifying agent the modifying agent being an organic compound
- A61K47/55—Medicinal preparations characterised by the non-active ingredients used, e.g. carriers or inert additives; Targeting or modifying agents chemically bound to the active ingredient the non-active ingredient being chemically bound to the active ingredient, e.g. polymer-drug conjugates the non-active ingredient being a modifying agent the modifying agent being an organic compound the modifying agent being also a pharmacologically or therapeutically active agent, i.e. the entire conjugate being a codrug
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P35/00—Antineoplastic agents
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D471/00—Heterocyclic compounds containing nitrogen atoms as the only ring hetero atoms in the condensed system, at least one ring being a six-membered ring with one nitrogen atom, not provided for by groups C07D451/00 - C07D463/00
- C07D471/02—Heterocyclic compounds containing nitrogen atoms as the only ring hetero atoms in the condensed system, at least one ring being a six-membered ring with one nitrogen atom, not provided for by groups C07D451/00 - C07D463/00 in which the condensed system contains two hetero rings
- C07D471/10—Spiro-condensed systems
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D487/00—Heterocyclic compounds containing nitrogen atoms as the only ring hetero atoms in the condensed system, not provided for by groups C07D451/00 - C07D477/00
- C07D487/02—Heterocyclic compounds containing nitrogen atoms as the only ring hetero atoms in the condensed system, not provided for by groups C07D451/00 - C07D477/00 in which the condensed system contains two hetero rings
- C07D487/04—Ortho-condensed systems
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D487/00—Heterocyclic compounds containing nitrogen atoms as the only ring hetero atoms in the condensed system, not provided for by groups C07D451/00 - C07D477/00
- C07D487/02—Heterocyclic compounds containing nitrogen atoms as the only ring hetero atoms in the condensed system, not provided for by groups C07D451/00 - C07D477/00 in which the condensed system contains two hetero rings
- C07D487/08—Bridged systems
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D487/00—Heterocyclic compounds containing nitrogen atoms as the only ring hetero atoms in the condensed system, not provided for by groups C07D451/00 - C07D477/00
- C07D487/02—Heterocyclic compounds containing nitrogen atoms as the only ring hetero atoms in the condensed system, not provided for by groups C07D451/00 - C07D477/00 in which the condensed system contains two hetero rings
- C07D487/10—Spiro-condensed systems
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D498/00—Heterocyclic compounds containing in the condensed system at least one hetero ring having nitrogen and oxygen atoms as the only ring hetero atoms
- C07D498/02—Heterocyclic compounds containing in the condensed system at least one hetero ring having nitrogen and oxygen atoms as the only ring hetero atoms in which the condensed system contains two hetero rings
- C07D498/10—Spiro-condensed systems
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D519/00—Heterocyclic compounds containing more than one system of two or more relevant hetero rings condensed among themselves or condensed with a common carbocyclic ring system not provided for in groups C07D453/00 or C07D455/00
Landscapes
- Chemical & Material Sciences (AREA)
- Organic Chemistry (AREA)
- Health & Medical Sciences (AREA)
- General Health & Medical Sciences (AREA)
- Veterinary Medicine (AREA)
- Medicinal Chemistry (AREA)
- Bioinformatics & Cheminformatics (AREA)
- Pharmacology & Pharmacy (AREA)
- Life Sciences & Earth Sciences (AREA)
- Animal Behavior & Ethology (AREA)
- Engineering & Computer Science (AREA)
- Public Health (AREA)
- General Chemical & Material Sciences (AREA)
- Chemical Kinetics & Catalysis (AREA)
- Nuclear Medicine, Radiotherapy & Molecular Imaging (AREA)
- Epidemiology (AREA)
- Nitrogen Condensed Heterocyclic Rings (AREA)
- Pharmaceuticals Containing Other Organic And Inorganic Compounds (AREA)
- Medicinal Preparation (AREA)
- Medicines That Contain Protein Lipid Enzymes And Other Medicines (AREA)
- Plural Heterocyclic Compounds (AREA)
- Nitrogen And Oxygen Or Sulfur-Condensed Heterocyclic Ring Systems (AREA)
- Peptides Or Proteins (AREA)
Description
関連出願の相互参照
本出願は、2020年10月1日に出願され、BRM TARGETING COMPOUNDS AND ASSOCIATED METHODS OF USEと題する米国非仮特許出願第16/590329号(これは、2019年4月1日に出願され、BRM TARGETING COMPOUNDS AND ASSOCIATED METHODS OF USEと題する米国非仮出願第16/372345号の一部継続である)に対する優先権を主張し、これは、2018年4月1日に出願され、BRM TARGETING PROTAC COMPOUNDS AND ASSOCIATED METHODS OF USEと題する米国仮特許出願第62/651,186号、および 2019年1月28日に出願され、BRM TARGETING PROTAC COMPOUNDS AND ASSOCIATED METHODS OF USEと題する米国仮特許出願第62/797,754号に対する優先権およびその利益を主張し、これらの各々は、参照によりそれらの全体が本明細書に組み込まれる。
CROSS-REFERENCE TO RELATED APPLICATIONS This application claims priority to U.S. Non-provisional Patent Application No. 16/590,329, filed October 1, 2020, entitled BRM TARGETING COMPOUNDS AND ASSOCIATED METHODS OF USE, which is a continuation-in-part of U.S. Non-provisional Application No. 16/372,345, filed April 1, 2019, entitled BRM TARGETING COMPOUNDS AND ASSOCIATED METHODS OF USE, which is a continuation-in-part of U.S. Non-provisional Application No. 16/372,345, filed April 1, 2018, entitled BRM TARGETING PROTAC COMPOUNDS AND ASSOCIATED METHODS OF USE. No. 62/651,186, filed Jan. 28, 2019, entitled BRM TARGETING PROTAC COMPOUNDS AND ASSOCIATED METHODS OF USE, and claims priority to and the benefit of U.S. Provisional Patent Application No. 62/797,754, filed Jan. 28, 2019, entitled BRM TARGETING PROTAC COMPOUNDS AND ASSOCIATED METHODS OF USE, each of which is incorporated herein by reference in its entirety.
本明細書は、標的タンパク質結合部分およびE3ユビキチンリガーゼ結合部分を含む二官能性化合物、ならびにその関連する使用方法を提供する。二官能性化合物は、標的化ユビキチン化の調節因子として、特に、本開示による二官能性化合物によって分解および/またはあるいは阻害される、スイッチ/スクロース非発酵性(SWI/SNF)関連、マトリックス関連、アクチン依存性調節因子のクロマチン、サブファミリーA、メンバー2(SMARCA2)(すなわち、BRAHMAまたはBRM)に関して有用である。 The present disclosure provides bifunctional compounds comprising a target protein binding portion and an E3 ubiquitin ligase binding portion, and related methods of use thereof. The bifunctional compounds are useful as regulators of targeted ubiquitination, particularly with respect to switch/sucrose non-fermenting (SWI/SNF)-associated, matrix-associated, actin-dependent regulator of chromatin, subfamily A, member 2 (SMARCA2) (i.e., BRAHMA or BRM), which is degraded and/or alternatively inhibited by the bifunctional compounds according to the present disclosure.
ほとんどの低分子薬剤は、酵素または受容体に密接に、および明確に定められたポケットにおいて結合する。一方で、低分子を使用してタンパク質-タンパク質相互作用を標的とすることは、タンパク質の接触表面が大きいこと、および関与するのが浅い溝状または平坦なインターフェースであるため困難であることが良く知られている。E3ユビキチンリガーゼ(そのうちの数百がヒトにおいて公知である)は、ユビキチン化に基質特異性を与え、したがって、特定のタンパク質基質に対する特異性があるため、一般的ななプロテアソームの阻害剤よりもさらに魅力的な治療標的である。E3リガーゼのリガンドの開発は、タンパク質-タンパク質の相互作用を破壊せざるを得ないという事実があるために部分的には困難であることが判明している。しかしながら、近年の開発によりこれらのリガーゼに結合する特定のリガンドがもたらされた。例えば、最初の低分子E3リガーゼ阻害剤であるナトリンの発見以来、E3リガーゼを標的とするさらなる化合物が報告されているが、未だこの分野は未開発である部分が多い。例えば、第1の小分子E3リガーゼマウス二重微小染色体2相同体(MDM2)阻害剤であるナトリンの発見以来、追加の化合物が標的MDM2(すなわち、ヒト二重微小染色体2またはHDM2)E3リガーゼ(J.Di,et al.Current Cancer Drug Targets(2011),11(8),987-994)を標的化することが報告されている。 Most small molecule drugs bind enzymes or receptors tightly and in well-defined pockets. On the other hand, targeting protein-protein interactions using small molecules is notoriously difficult due to the large protein contact surfaces and shallow groove-like or flat interfaces involved. E3 ubiquitin ligases (hundreds of which are known in humans) confer substrate specificity to ubiquitination and are therefore more attractive therapeutic targets than general proteasome inhibitors due to their specificity for specific protein substrates. Development of ligands for E3 ligases has proven difficult in part due to the fact that they are forced to disrupt protein-protein interactions. However, recent developments have provided specific ligands that bind to these ligases. For example, since the discovery of the first small molecule E3 ligase inhibitor, nutrin, additional compounds targeting E3 ligases have been reported, but the field is still largely unexplored. For example, since the discovery of nutrin, the first small molecule E3 ligase mouse double minute 2 homologue (MDM2) inhibitor, additional compounds have been reported to target MDM2 (i.e., human double minute 2 or HDM2) E3 ligase (J. Di, et al. Current Cancer Drug Targets (2011), 11(8), 987-994).
魅力的な治療可能性のあるE3リガーゼの1つが、フォンヒッペル-リンドウ(von Hippel-Lindau(VHL))腫瘍抑制因子であり、これはE3リガーゼ複合体VCBの基質認識サブユニットであり、この複合体はさらにエロンギン(elongin)BおよびC、Cul2ならびにRbx1からなる。VHLの一次基質は、低酸素レベルに応じて血管内皮細胞増殖因子VEGFおよび赤血球誘発サイトカインエリスロポエチンなどの遺伝子を上方制御する転写因子である、低酸素誘導因子1α(HIF-1α)である。E3リガーゼの基質認識サブユニットに対するフォンヒッペル-リンドウ(VHL)の最初の低分子リガンドが作製され、結晶構造が取得され、この化合物がVHLの主要基質である転写因子HIF-1αの結合様式を模倣することが確認された。 One E3 ligase with attractive therapeutic potential is the von Hippel-Lindau (VHL) tumor suppressor, which is the substrate recognition subunit of the E3 ligase complex VCB, which further consists of elongins B and C, Cul2, and Rbx1. The primary substrate of VHL is hypoxia-inducible factor 1α (HIF-1α), a transcription factor that upregulates genes such as the vascular endothelial growth factor VEGF and the erythropoietin-inducing cytokine in response to low oxygen levels. The first small molecule ligand of von Hippel-Lindau (VHL) to the substrate recognition subunit of E3 ligase has been generated and a crystal structure obtained, confirming that this compound mimics the binding mode of the transcription factor HIF-1α, the primary substrate of VHL.
米国特許出願公開第2015/0291562号および同第2014/0356322号(参照により本明細書に組み込まれる)に記載されるものなどの二官能性化合物は、分解のためにE3ユビキインリガーゼに内因性タンパク質を動員するように機能する。特に当該特許公開は、二官能性またはタンパク溶解性の標的化キメラ(PROTAC:proteolysis targeting chimeric)化合物を記載しており、当該化合物は様々なポリペプチドおよび他のタンパク質の標的ユビキチン化の調節因子としての有用性が見いだされ、それらは当該二官能性化合物によって分解および/または別手段により阻害される。 Bifunctional compounds such as those described in U.S. Patent Application Publication Nos. 2015/0291562 and 2014/0356322 (herein incorporated by reference) function to recruit endogenous proteins to E3 ubiquiin ligases for degradation. In particular, the patent publications describe bifunctional or proteolysis targeting chimeric (PROTAC) compounds that find utility as regulators of targeted ubiquitination of various polypeptides and other proteins, which are degraded and/or otherwise inhibited by the bifunctional compounds.
スイッチ/スクロース非発酵性(SWI/SNF)は、2つの相互排他的なヘリカーゼ/ATPase触媒サブユニットであるSWI/SNF関連、マトリックス関連、アクチン依存性調節因子のクロマチン、サブファミリーA、メンバー2(SMARCA2、BRAHMA、またはBRM)およびSWI/SNF関連、マトリックス関連、アクチン依存性調節因子のクロマチン、サブファミリーA、メンバー4(SMARCA4またはBRG1)の活性によって色素構造を調節する、多サブユニット複合体である。コアサブユニットおよび調節サブユニットはATP加水分解をヒストン-DNA接触の摂動に結合し、それによって遺伝子活性および抑制を促進する転写因子および同族DNA要素への接続点を提供する。 Switch/sucrose nonfermenting (SWI/SNF) is a multisubunit complex that regulates chromatin structure through the activity of two mutually exclusive helicase/ATPase catalytic subunits, SWI/SNF-related, matrix-associated, actin-dependent regulator of chromatin, subfamily A, member 2 (SMARCA2, BRAHMA, or BRM) and SWI/SNF-related, matrix-associated, actin-dependent regulator of chromatin, subfamily A, member 4 (SMARCA4 or BRG1). The core and regulatory subunits couple ATP hydrolysis to perturbations of histone-DNA contacts, thereby providing attachment points to transcription factors and cognate DNA elements that promote gene activity and repression.
ラブドイド腫瘍、女性がん(卵巣がん、子宮がん、子宮頸がん、および子宮内膜がんを含む)、肺腺がん、胃腺がん、黒色腫瘍、食道がん、および腎明細胞がんにおいて観察される最も高頻度の変異を有するすべてのがんのほぼ20%において、20個のカノニカルSWI/SNFサブユニットをコードする遺伝子における変異が観察される。高程度の相同性、およびそれらの推定される重複機能を有するにもかかわらず、SMARCA2およびSMARCA4は、がんにおいて異なる役割を有すると報告されている。例えば、SMARCA4は、原発性腫瘍において頻繁に変異し、一方、SMARCA2の不活性化は、腫瘍形成において稀である。実際、様々な種類のがんは、例えば、肺がん(例えば、非小細胞肺がん)を含む、SMARCA4関連(例えば、SMARCA4変異、または発現の欠如などのSMARCA4欠損を有するがん)であることが示されている。 Mutations in genes encoding the 20 canonical SWI/SNF subunits are observed in nearly 20% of all cancers, with the most frequent mutations observed in rhabdoid tumors, female cancers (including ovarian, uterine, cervical, and endometrial cancers), lung adenocarcinoma, gastric adenocarcinoma, melanoma, esophageal cancer, and renal clear cell carcinoma. Despite having a high degree of homology and their presumed overlapping functions, SMARCA2 and SMARCA4 have been reported to have different roles in cancer. For example, SMARCA4 is frequently mutated in primary tumors, while inactivation of SMARCA2 is rare in tumorigenesis. Indeed, various types of cancer have been shown to be SMARCA4-associated (e.g., cancers with SMARCA4 mutations or SMARCA4 defects, such as lack of expression), including, for example, lung cancer (e.g., non-small cell lung cancer).
SMARCA2は、SMARCA4関連または変異型がん細胞株においてもっとも必須とされる遺伝子のうちの1つとして実証されている。これは、SMARCA4欠損患者集団または細胞がSMARCA2活性にのみ依存するためであり、すなわち、SMARCA2が複合体中により大きく組み込まれてSMARCA4欠損を補填するからである。したがって、SMARCA2は、SMARCA4関連/欠損がんにおいて標的化され得る。細胞死をもたらす2つ(以上)の遺伝子の発現の欠如の共起は、合成致死として既知である。したがって、合成致死は、ある特定のSMARCA2/SMARCA4関連がんの治療において活用することができる。 SMARCA2 has been demonstrated as one of the most essential genes in SMARCA4-associated or mutant cancer cell lines. This is because SMARCA4-deficient patient populations or cells are solely dependent on SMARCA2 activity, i.e., SMARCA2 is more highly incorporated into the complex to compensate for the SMARCA4 deficiency. Therefore, SMARCA2 can be targeted in SMARCA4-associated/deficient cancers. The co-occurrence of the lack of expression of two (or more) genes resulting in cell death is known as synthetic lethality. Thus, synthetic lethality can be exploited in the treatment of certain SMARCA2/SMARCA4-associated cancers.
SMARCA2(すなわち、BRAHMAまたはBRM)を阻害または分解することによって治療可能である疾患に対する有効な治療法が継続的に必要とされている。しかしながら、非特異的な効果、ならびにSMARCA2を標的化および調節することができないことは、効果的な治療の開発の障害であり続ける。したがって、SMARCA2を標的化し、VHLの基質特異性を活用または増強する小分子治療剤は、非常に有用であろう。 There is a continuing need for effective therapies for diseases treatable by inhibiting or degrading SMARCA2 (i.e., BRAHMA or BRM). However, nonspecific effects and the inability to target and regulate SMARCA2 continue to be obstacles to the development of effective treatments. Thus, small molecule therapeutics that target SMARCA2 and exploit or enhance the substrate specificity of VHL would be highly useful.
本開示は、内因性タンパク質をE3ユビキチンリガーゼへとリクルートし、分解するよう機能する二官能性化合物、およびその使用方法を記載するものである。特に本開示は二官能性またはタンパク溶解性の標的化キメラ(proteolysis targeting chimeric、PROTAC)化合物を提供するものであり、当該化合物は様々なポリペプチドおよび他のタンパク質の標的ユビキチン化の調節因子としての有用性が見いだされ、それらポリペプチドおよび他のタンパク質は本明細書に記載される二官能性化合物によって分解および/または別手段により阻害される。本明細書に提供される化合物の利点は、広範な薬理活性がある可能性があり、事実上、すべてのタンパク質種またはファミリーからの標的ポリペプチドの分解/阻害と調和する。加えて、本明細書は、がん、例えば、肺がんまたは非小細胞肺がんなどのSMARCA4関連/欠損がんなどの疾患状態の治療または改善のために有効量の本明細書に記載される化合物を使用する方法を提供する。 The present disclosure describes bifunctional compounds that function to recruit endogenous proteins to E3 ubiquitin ligases for degradation, and methods of use thereof. In particular, the present disclosure provides bifunctional or proteolytic targeting chimeric (PROTAC) compounds that find utility as regulators of targeted ubiquitination of various polypeptides and other proteins that are degraded and/or otherwise inhibited by the bifunctional compounds described herein. The advantage of the compounds provided herein is that they may have a broad range of pharmacological activity, consistent with the degradation/inhibition of target polypeptides from virtually any protein species or family. In addition, the present disclosure provides methods of using an effective amount of the compounds described herein for the treatment or amelioration of disease conditions, such as cancer, e.g., SMARCA4-associated/deficient cancers, such as lung cancer or non-small cell lung cancer.
したがって、1つの態様において本開示は二官能性化合物またはPROTAC化合物を提供するものであり、当該化合物は、E3ユビキチンリガーゼ結合部分(すなわち、E3ユビキチンリガーゼに対するリガンド、または「ULM」基)と、標的タンパク質に結合する部分(すなわちタンパク質/ポリペプチド標的化リガンド、または「PTM」基)を含み、それにより当該標的のタンパク質/ポリペプチドはユビキチンリガーゼに近接して配置され、当該タンパク質の分解(および阻害)作用が発揮される。好ましい実施形態では、ULM(ユビキチン化リガーゼ調節因子)は、フォンヒッペル・リンダウE3ユビキチンリガーゼ(VHL)結合部分(VLM)であり得る。例えば、二官能性化合物の構造を以下のように示すことができる:
本明細書に例示されるPTMおよびULM部分のそれぞれの位置、ならびにそれらの数は、例としてのみ提供され、化合物をいかなる方法でも制限することを意図するものではない。当業者によって理解されるように、本明細書に記載される二官能性化合物は、それぞれの官能性部分の数および位置が所望のとおり変化することができるように合成することができる。 The respective positions of the PTM and ULM moieties, as well as their numbers, illustrated herein are provided by way of example only and are not intended to limit the compounds in any manner. As will be appreciated by one of skill in the art, the bifunctional compounds described herein can be synthesized such that the number and positions of the respective functional moieties can be varied as desired.
特定の実施形態では、二官能性化合物は化学リンカー(「L」)をさらに含む。本実施例において、二官能性化合物の構造は以下のように示すことができる:
例えば、二官能性化合物の構造を以下のように示すことができる:
特定の実施形態では、本明細書に記載の化合物は、ULM、複数のPTM、複数の化学リンカー、またはそれらの組み合わせから独立して選択される複数を含む。 In certain embodiments, the compounds described herein include a plurality of independently selected ULMs, a plurality of PTMs, a plurality of chemical linkers, or a combination thereof.
追加的実施形態では、VLMはヒドロキシプロリンまたはその誘導体であり得る。さらに、他の企図されるVLMは、上記に考察されるように、その全体が本明細書に組み込まれる米国特許出願公開第2014/03022523号に含まれる。 In additional embodiments, the VLM can be hydroxyproline or a derivative thereof. Additionally, other contemplated VLMs are included in U.S. Patent Application Publication No. 2014/03022523, which is incorporated herein in its entirety, as discussed above.
特定の実施形態では、「L」は結合である。追加的実施形態では、リンカー「L」は、1~20の範囲の直線状の非水素原子数を有するコネクターである。コネクターの「L」は、例えばエーテル、アミド、アルカン、アルケン、アルキン、ケトン、ヒドロキシル、カルボン酸、チオエーテル、スルホキシド、およびスルホンなどの官能基を含み得るが、これらに限定されない。リンカーは、芳香族、芳香族複素環式、環式、二環式、および三環式の部分を含み得る。Cl、F、BrおよびIなどのハロゲンとの置換が、リンカーに含まれてもよい。フッ素置換の場合、単一または複数のフッ素が含まれてもよい。 In certain embodiments, "L" is a bond. In additional embodiments, the linker "L" is a connector having a linear number of non-hydrogen atoms ranging from 1 to 20. The connector "L" may include functional groups such as, but not limited to, ether, amide, alkane, alkene, alkyne, ketone, hydroxyl, carboxylic acid, thioether, sulfoxide, and sulfone. The linker may include aromatic, heteroaromatic, cyclic, bicyclic, and tricyclic moieties. Substitution with halogens such as Cl, F, Br, and I may be included in the linker. In the case of fluorine substitution, single or multiple fluorines may be included.
特定の実施形態では、VLMは、トランス-3-ヒドロキシプロリンの誘導体であり、ここでトランス-3-ヒドロキシプロリン中の窒素とカルボン酸の両方が、アミドとして官能性がある。 In certain embodiments, the VLM is a derivative of trans-3-hydroxyproline, where both the nitrogen and the carboxylic acid in trans-3-hydroxyproline are functionalized as amides.
追加の態様において本明細書は、本明細書に記載の化合物またはその塩型の有効量、および薬学的に許容可能な担体を含む治療用組成物を提供する。治療用組成物は、患者または対象、例えばヒトなどの動物においてタンパク質の分解および/または阻害を調節し、当該分解されたおよび/または阻害されたタンパク質を介して調節される病態もしくは状態の治療または改善に使用することができる。ある特定の実施形態では、本明細書に記載される治療組成物を使用して、疾患、例えば、肺がん(例えば、非小細胞肺がん)などのがん(SWI/SNF関連がん、SMARCA4変異を有するがん、SMARCA4欠損を有するがん、またはそれらの組み合わせのうちの少なくとも1つを含む)の治療または改善のために、目的のタンパク質の分解をもたらし得る。さらに別の態様において本開示は、細胞中の標的タンパク質をユビキチン化/分解する方法を提供する。ある特定の実施形態では、本方法は、本明細書に別途記載されるように、好ましくはリンカー部分を介して結合されるVLMを含む、本明細書に記載される二官能性化合物を投与することを含み、VLMは、分解のためにタンパク質を標的化するために、リンカーを介してPTMに結合される。標的タンパク質の分解は、その標的タンパク質がE3ユビキチンリガーゼに近接して配置されたときに生じ、したがって、標的タンパク質の分解/標的タンパク質の効果の阻害、およびタンパク質レベルの制御がもたらされる。本開示によりもたらされるタンパク質レベルの制御は病態または状態の治療を提供するものであり、これは患者細胞中のそのタンパク質のレベルを低下させることによって標的タンパク質を介して調節される。 In an additional aspect, the present disclosure provides a therapeutic composition comprising an effective amount of a compound or salt form thereof described herein and a pharma- ceutically acceptable carrier. The therapeutic composition can be used to modulate protein degradation and/or inhibition in a patient or subject, e.g., an animal, such as a human, and to treat or ameliorate a disease state or condition modulated via the degraded and/or inhibited protein. In certain embodiments, the therapeutic composition described herein can be used to effect degradation of a protein of interest for the treatment or amelioration of a disease, e.g., cancer, such as lung cancer (e.g., non-small cell lung cancer), including at least one of SWI/SNF-associated cancer, cancer with SMARCA4 mutation, cancer with SMARCA4 deficiency, or a combination thereof. In yet another aspect, the present disclosure provides a method of ubiquitinating/degrading a target protein in a cell. In certain embodiments, the method includes administering a bifunctional compound described herein, preferably comprising a VLM linked via a linker moiety, as described elsewhere herein, wherein the VLM is linked to a PTM via a linker to target the protein for degradation. Degradation of a target protein occurs when the target protein is placed in close proximity to an E3 ubiquitin ligase, thus resulting in degradation of the target protein/inhibition of the effect of the target protein and control of protein levels. The control of protein levels provided by the present disclosure provides treatment of a disease state or condition that is regulated via the target protein by reducing the level of that protein in the patient's cells.
さらに別の態様において本明細書は、例えばヒトなどの動物などの対象または患者において、疾患、障害もしくはその症状を治療または改善する方法を提供するものであり、当該方法は、本明細書に記載の化合物またはその塩型の治療有効量などの有効量、および薬学的に許容可能な担体を含む組成物をその必要のある対象に投与することを含み、組成物は、対象において疾患もしくは障害またはその症状の治療または改善に有効である。 In yet another aspect, the present specification provides a method of treating or ameliorating a disease, disorder, or a symptom thereof in a subject or patient, e.g., an animal, e.g., a human, the method comprising administering to a subject in need thereof a composition comprising an effective amount, e.g., a therapeutically effective amount, of a compound, or a salt form thereof, as described herein, and a pharma- ceutically acceptable carrier, wherein the composition is effective to treat or ameliorate the disease or disorder, or a symptom thereof, in the subject.
別の態様において本明細書は、本開示による化合物を使用して生物的システムにおいて対象タンパク質の分解効果を特定する方法を提供する。 In another aspect, the present specification provides a method for identifying the degradative effect of a protein of interest in a biological system using a compound according to the present disclosure.
有用性に関する前述の全般的な記載は、例示の目的でのみ提示するものであり、本開示および添付の請求の範囲を限定することは意図されない。本開示の組成物、方法、およびプロセスに関連する追加の目的および利点は、請求の範囲、詳細な説明、および実施例を鑑みることで当分野の当業者に理解されるであろう。例えば本開示の様々な態様および実施形態は多くの組み合わせで使用することができ、それらすべてが本明細書により明示的に予期される。これらの追加的な態様および実施形態は、本開示の範囲内に明示的に含まれる。本開示の背景を解説するために、および特定の場合には実施に関して追加の詳細を提供するために本明細書において使用される公表文献および他の資料は、参照により組み込まれる。 The foregoing general description of utility is presented for illustrative purposes only and is not intended to limit the scope of the disclosure and the appended claims. Additional objects and advantages associated with the compositions, methods, and processes of the present disclosure will be understood by those of ordinary skill in the art in view of the claims, detailed description, and examples. For example, the various aspects and embodiments of the present disclosure can be used in many combinations, all of which are expressly contemplated by this specification. These additional aspects and embodiments are expressly included within the scope of the present disclosure. Publications and other materials used herein to describe the background of the disclosure and, in certain cases, to provide additional details regarding the implementation are incorporated by reference.
本明細書の一部に組み込まれ、その一部を形成する添付図面は、本開示のいくつかの実施形態を図示し、本明細書の記載とともに本開示の原理を説明する役割を果たす。図面は、本開示の実施形態の解説の目的のためのみであり、本開示を限定すると解釈されるものではない。本開示のさらなる目的、特徴および利点は、本開示の例示的実施形態を示す添付図面と併せ、以下の詳細な説明から明白となるであろう。 The accompanying drawings, which are incorporated in and form a part of this specification, illustrate several embodiments of the present disclosure and, together with the description herein, serve to explain the principles of the present disclosure. The drawings are for the purpose of illustrating embodiments of the present disclosure only and are not to be construed as limiting the present disclosure. Further objects, features and advantages of the present disclosure will become apparent from the following detailed description taken in conjunction with the accompanying drawings showing exemplary embodiments of the present disclosure.
以下は、本開示の実施において当業者を支援するために提供される詳細な説明である。当分野の当業者であれば、本開示の趣旨または範囲から逸脱することなく、本明細書に記載の実施形態の修正および変更を行い得る。本明細書において言及されるすべての公表文献、特許出願、特許、図面およびその他の参考文献は、参照によりその全体が明示的に組み込まれる。 The following is a detailed description provided to assist those skilled in the art in practicing the present disclosure. Those skilled in the art may make modifications and variations to the embodiments described herein without departing from the spirit or scope of the present disclosure. All publications, patent applications, patents, figures and other references mentioned herein are expressly incorporated by reference in their entirety.
E3ユビキチンリガーゼタンパク質と標的タンパク質とを結合する二官能性またはキメラ構築物によって、一度E3ユビキチンリガーゼタンパク質と標的タンパク質とが近接して配置されると、E3ユビキチンリガーゼタンパク質(例えば、フォンヒッペル・リンダウE3ユビキチンリガーゼ(VHL))が標的タンパク質をユビキチン化するという驚くべき予期せぬ発見に関連する、組成物および方法が本明細書に記載される。したがって、本開示は、タンパク質標的結合部分(PTM)に結合したE3ユビキチンリガーゼ結合部分(ULM)を含むような化合物および組成物を提供するものであり、これにより選択標的タンパク質のユビキチン化が生じ、それに伴いプロテアソームによる標的タンパク質の分解が導かれる(図1を参照)。本開示はまた、組成物のライブラリおよびその使用を提供する。 Described herein are compositions and methods related to the surprising and unexpected discovery that an E3 ubiquitin ligase protein (e.g., von Hippel-Lindau E3 ubiquitin ligase (VHL)) ubiquitinates a target protein once the two proteins are brought into close proximity via a bifunctional or chimeric construct that links the E3 ubiquitin ligase protein to the target protein. Accordingly, the present disclosure provides compounds and compositions that include an E3 ubiquitin ligase binding moiety (ULM) linked to a protein target binding moiety (PTM), which results in ubiquitination of a selected target protein and associated degradation of the target protein by the proteasome (see FIG. 1). The present disclosure also provides a library of compositions and uses thereof.
ある特定の態様では、本開示は、VHLなどのユビキチンリガーゼに結合することができるリガンド、例えば、小分子リガンド(すなわち、2,000、1,000、500、または200ダルトン未満の分子量を有する)を含む化合物を提供する。化合物は、標的タンパク質をユビキチンリガーゼに近接して配置させ、当該タンパク質の分解(および/または阻害)を生じさせるようなやり方で標的タンパク質に結合することができる部分も含む。上記に加えて低分子とは、分子が非ぺプチジルであることを意味し得る。すなわち例えば4、3、または2個よりも少ないアミノ酸を含むなど、多くの場合ペプチドとはみなされない。本明細書によると、PTM、ULM、またはPROTAC分子は低分子であってもよい。 In certain aspects, the disclosure provides compounds that include a ligand, e.g., a small molecule ligand (i.e., having a molecular weight of less than 2,000, 1,000, 500, or 200 daltons), that can bind to a ubiquitin ligase, such as VHL. The compound also includes a moiety that can bind to a target protein in a manner that places the target protein in proximity to a ubiquitin ligase, resulting in degradation (and/or inhibition) of the protein. In addition to the above, small molecule can mean that the molecule is non-peptidyl, i.e., often not considered a peptide, e.g., containing fewer than 4, 3, or 2 amino acids. In accordance with the present specification, a PTM, ULM, or PROTAC molecule can be a small molecule.
別途定義されない限り、本明細書で使用されるすべての技術用語および科学的用語は、本開示が属する分野の当業者によって一般的に理解される意味と同じ意味を有する。本明細書で使用される専門用語は、特定の実施形態の解説のみを目的としており、本開示を限定することは意図されていない。 Unless otherwise defined, all technical and scientific terms used herein have the same meaning as commonly understood by one of ordinary skill in the art to which this disclosure belongs. The terminology used herein is for the purpose of describing particular embodiments only and is not intended to be limiting of the disclosure.
値の範囲が提供される場合、文脈により明白に別段の指定が無い限り(例えばある数の炭素原子を含有する基の場合、当該範囲内におさまる各炭素原子数が提供される)、その範囲および任意の他の指定範囲の上限と下限の間の、下限単位の十分の一までの各介在値、またはその指定範囲中の介在値が本発明範囲内に包含されることを理解されたい。これらのより小さな範囲の上限および下限が独立してより小さな範囲に含まれてもよく、これも本開示内に包含され、指定範囲中の任意の具体的に除外される境界値となる。指定範囲が境界値の一方または両方を含む場合、それら含まれる境界値のいずれか、両方を除外する範囲も本開示に含まれる。 When a range of values is provided, it is to be understood that each intervening value between the upper and lower limits of that range and any other specified range, to one-tenth of the unit of the lower limit, or in that specified range, is encompassed within the scope of the invention, unless the context clearly dictates otherwise (e.g., in the case of a group containing a certain number of carbon atoms, each number of carbon atoms falling within the range is provided). The upper and lower limits of these smaller ranges may be independently included in the smaller ranges, which are also encompassed within the disclosure, as are any specifically excluded limits in the specified ranges. When a specified range includes one or both of the limits, ranges excluding either or both of those included limits are also included in the disclosure.
以下の用語は、本開示を説明するために使用される。本明細書において用語が具体的に定義されていない場合、その用語は、本開示の説明においてその使用の分若で当該用語を適用する当業者により当分野において認識されている意味が与えられる。 The following terms are used to describe this disclosure. If a term is not specifically defined herein, the term is given its art-recognized meaning by those of ordinary skill in the art who apply the term to the extent that it is used in describing this disclosure.
本明細書において使用される場合、「a」および「an」という冠詞は、文脈により明白に別段の示唆が無い限り、当該冠詞の文法的客体のうちの1つまたは1つ以上(すなわち少なくとも1つ)を指すように本明細書において使用される。一例として、「要素」は1つの要素または2つ以上の要素を意味する。 As used herein, the articles "a" and "an" are used herein to refer to one or to more than one (i.e., to at least one) of the grammatical object of the article, unless the context clearly indicates otherwise. By way of example, "an element" means one element or more than one element.
本明細書および請求の範囲において本明細書に使用される場合、「および/または」という語句は、そのように結合される要素のうちの「いずれか、または両方」を意味すると理解されたい。すなわち、一部の例では要素は結合して存在し、他の例では結合せずに存在する。「および/または」を用いて列記された複数の要素は、同じように解釈されるべきである。すなわち、要素の「1つ以上」がそのように結合されている。「および/または」条項により具体的に特定された要素以外の他の要素が、それら具体的に特定された要素との関連性の有無に関係なく、任意に存在し得る。したがって、非限定的な例として、「Aおよび/またはB」への言及は、「含む」などのオープンエンド言語と併用される場合、一実施形態では、Aのみ(任意にB以外の要素を含む)、別の実施形態では、Bのみ(任意にA以外の要素を含む)、さらに別の実施形態では、AおよびBの両方(任意に他の要素を含む)などを指すことができる。 As used herein in the specification and claims, the phrase "and/or" should be understood to mean "either or both" of the elements so conjoined. That is, in some instances the elements are conjointly present and in other instances they are non-conjoined. Multiple elements listed with "and/or" should be interpreted in the same manner, that is, "one or more" of the elements are so conjoined. Other elements other than the elements specifically identified by the "and/or" clause may optionally be present, whether or not related to those specifically identified elements. Thus, as a non-limiting example, a reference to "A and/or B," when used in conjunction with open-ended language such as "comprising," may refer in one embodiment to only A (optionally including elements other than B), in another embodiment to only B (optionally including elements other than A), in yet another embodiment to both A and B (optionally including other elements), etc.
本明細書および請求の範囲において本明細書で使用される場合、「または」は、上記に定義される「および/または」と同じ意味を有すると理解されるべきである。例えば、リスト内の項目を分離するときに、「または」または「および/または」は包括的なものとして解釈されるものとする。すなわち、多くの要素、または要素のリストのうちの少なくとも1つを含むが、2つ以上も含み、そして任意で列記されていない追加の項目も含む。例えば「~の内のただ1つ」、または「~の内の正確に1つ」、または請求項において使用される場合には「~からなる」など、逆を明確に示唆される用語のみが、多くの要素、または要素のリストのうちの正確に1つの要素の含有を指す。概して、本明細書に使用される場合、「または」という用語は、例えば「いずれか」、「~のうちの1つ」、「~のうちの1つのみ」、または「~のうちの正確に1つ」などの排他的な用語が先行する場合にのみ排他的な選択肢(すなわち「1つまたはその他であるが、両方ではない」)を示すものと解釈されるべきである。 As used herein in the specification and claims, "or" should be understood to have the same meaning as "and/or" as defined above. For example, when separating items in a list, "or" or "and/or" shall be interpreted as inclusive; that is, including at least one of a number or list of elements, but also including two or more, and optionally including additional items not listed. Only terms that clearly suggest the contrary, such as "only one of," or "exactly one of," or, when used in the claims, "consisting of," refer to the inclusion of exactly one element of a number or list of elements. In general, as used herein, the term "or" should be interpreted to indicate exclusive alternatives (i.e., "one or the other, but not both") only when preceded by an exclusive term, such as, for example, "either," "one of," "only one of," or "exactly one of."
請求の範囲、ならびに上述の明細書において、「含む(comprising)」、「含有する(including)」、「担持する(carrying)」、「有する(having)」「含有する(containing)」、「関与する(involving)」、「保持する(holding)」、「構成される(composed of)」などのすべての移行句は、非限定である、すなわちそれらを含むが限定されないことを意味すると理解されたい。「~からなる(consisting of)」および「本質的に~からなる(consisting essentially of)」という移行句のみ、それぞれ限定的または半限定的な移行句であるものとし、このことは米国特許審査基準のセクション2111.03に記載されている。 In the claims, as well as in the above specification, all transitional phrases such as "comprising," "including," "carrying," "having," "containing," "involving," "holding," "composed of," and the like, are to be understood to be open-ended, i.e., meaning including but not limited to. Only the transitional phrases "consisting of" and "consisting essentially of" shall be closed or semi-closed transitional phrases, respectively, as set forth in Section 2111.03 of the United States Patent Examining Guidelines.
本明細書において使用される場合、明細書および請求の範囲において、1つ以上の要素のリストに関し、「少なくとも1つ」という語句は、要素リスト中のいずれか1つ、または複数の要素から選択される少なくとも1つの要素を意味すると理解されるべきであるが、必ずしも要素リスト内に具体的に列記されるすべての要素のうちの少なくとも1つを含むものではなく、要素リスト中の要素の任意の組み合わせを除外するものではない。さらにこの定義は、「少なくとも1つ」という語句が指す要素リスト内で具体的に特定された要素以外にも、具体的に特定されたそれらの要素の関連性の有無にかかわらず、任意で要素が存在し得ることを許容する。したがって、非限定的な例として、「AおよびBのうちの少なくとも1つ」(または同等に、「AまたはBのうちの少なくとも1つ」、または同等に、「Aおよび/またはBのうちの少なくとも1つ」)は、一実施形態では、少なくとも1つの、任意に、2つ以上のAを含み、Bが存在しない(かつ、任意に、B以外の要素を含む)、別の実施形態では、少なくとも1つの、任意に、2つ以上のBを含み、Aが存在しない(かつ、任意に、A以外の要素を含む)、さらに別の実施形態では、少なくとも1つの、任意に、2つ以上のA、ならびに少なくとも1つの、任意に、2つ以上のBを含む(かつ、任意に、他の要素を含む)を指すことができる。 As used herein, in the specification and claims, the phrase "at least one" in reference to a list of one or more elements should be understood to mean at least one element selected from any one or more of the elements in the list of elements, but not necessarily including at least one of all elements specifically listed in the list of elements, and not excluding any combination of elements in the list of elements. Furthermore, this definition allows for the optional presence of elements other than those specifically identified in the list of elements to which the phrase "at least one" refers, regardless of the relevance of those specifically identified elements. Thus, as a non-limiting example, "at least one of A and B" (or, equivalently, "at least one of A or B" or, equivalently, "at least one of A and/or B") can refer in one embodiment to at least one, optionally, two or more A, and no B (and, optionally, including elements other than B); in another embodiment to at least one, optionally, two or more B, and no A (and, optionally, including elements other than A); and in yet another embodiment to at least one, optionally, two or more A, and at least one, optionally, two or more B (and, optionally, including other elements).
2つ以上の工程または動作を含む本明細書に記載の特定の方法において、方法の工程または動作の順序は、文脈より別段の示唆が無い限り、方法の工程または動作が列挙される順序に必ずしも限定されない。 In certain methods described herein that include more than one step or action, the order of the method steps or actions is not necessarily limited to the order in which the method steps or actions are recited, unless the context indicates otherwise.
「同時投与(co-administration)」および「同時に投与すること(co-administering)」または「併用療法(combination therapy)」という用語は、同時投与(2つ以上の治療薬の同時投与)と、治療薬が患者の体内にある程度、好ましくは効果量が同時に存在する間は時間を変えて投与すること(1つ以上の治療薬を、追加の治療薬の投与の時間とは異なる時間に投与すること)との両方を指す。特定の好ましい態様において、本明細書に記載の本化合物の1つ以上が、特に抗がん剤を含む、少なくとも1つの追加生物活性剤と併用して共投与される。特に好ましい態様において、化合物の共投与により、抗がん活性を含む相乗作用的な活性および/または治療がもたらされる。 The terms "co-administration" and "co-administering" or "combination therapy" refer to both simultaneous administration (administration of two or more therapeutic agents at the same time) and staggered administration (administration of one or more therapeutic agents at a time different from the time of administration of the additional therapeutic agent) while the therapeutic agents are simultaneously present in the patient's body in some, preferably effective, amount. In certain preferred embodiments, one or more of the present compounds described herein are co-administered in combination with at least one additional bioactive agent, including, in particular, anti-cancer agents. In particularly preferred embodiments, co-administration of the compounds results in synergistic activity and/or therapy, including anti-cancer activity.
本明細書に使用される場合、別段の示唆が無い限り、「化合物」という用語は、本明細書に開示される任意の具体的な化学的化合物を指し、互変異性体、位置異性体、幾何異性体、および適切な場合には光学異性体(鏡像異性体)およびその他の立体異性体(ジアステレオマー)を含む立体異性体、ならびに文脈で適切な場合にはそのプロドラッグおよび/または重水素化型を含む薬学的に許容可能な塩および誘導体を含む。予期される重水素化低分子は、薬剤分子中に含有される水素原子のうちの1つ以上が重水素で置換されている低分子である。 As used herein, unless otherwise indicated, the term "compound" refers to any specific chemical compound disclosed herein, including tautomers, positional isomers, geometric isomers, and, where appropriate, stereoisomers, including optical isomers (enantiomers) and other stereoisomers (diastereomers), as well as pharma- ceutically acceptable salts and derivatives thereof, including prodrugs and/or deuterated forms, where appropriate in the context. Contemplated deuterated small molecules are small molecules in which one or more of the hydrogen atoms contained in the drug molecule have been replaced with deuterium.
文脈中のその使用の内で、化合物という用語は概して単一の化合物を指すが、例えば本開示化合物の立体異性体、位置異性体および/または光学異性体(ラセミ混合物を含む)ならびに特定の鏡像異性体または特定の鏡像異性体を富化した混合物などの他の化合物を含んでもよい。当該用語は文脈において、投与を促進し、活性部位へ化合物を送達するよう改変された化合物のプロドラッグ型も指す。本化合物の記載において、多くの置換基、および特にそれと関連した変数が記載されていることに注意されたい。本明細書に記載される分子は、以下に概略されるように安定した化合物であることが当業者には理解される。結合が示されている場合、二重結合と一重結合の両方とも、示されている化合物、および価数の相互作用に関する公知のルールを背景として表され、または理解される。 Within its use in context, the term compound generally refers to a single compound, but may include other compounds, such as stereoisomers, regioisomers, and/or optical isomers (including racemic mixtures) of the disclosed compounds, as well as specific enantiomers or mixtures enriched for specific enantiomers. The term also refers in context to prodrug forms of compounds that have been modified to facilitate administration and deliver the compound to the site of activity. Note that in describing the compounds, many substituents and variables, particularly those associated therewith, are described. It will be understood by those of skill in the art that the molecules described herein are stable compounds, as outlined below. Where bonds are shown, both double and single bonds are depicted or understood in the context of the compounds shown and known rules regarding valency interactions.
「ユビキチンリガーゼ」という用語は、特定の基質タンパク質へのユビキチンの移送を促進し、その基質タンパク質を分解の標的とさせるタンパク質のファミリーを指す。例えば、E3ユビキチンリガーゼタンパク質は、単独で、またはE2ユビキチン共役酵素と組み合わせて、標的タンパク質上のリジンへのユビキチンの結合を引き起こし、その後、プロテアソームによる分解のために特定のタンパク質基質を標的化する。したがって、E2ユビキチン結合酵素との複合体で、または単独で、E3ユビキチンリガーゼは、標的とされるタンパク質へのユビキチンの移送に関与する。概して、ユビキチンリガーゼは、ポリユビキチン化に関与し、それにより第二のユビキチンが第一のユビキチンに付加され、第三のユビキチンが第二のユビキチンに付加される。ポリユビキチン化は、プロテアソーム分解に対してタンパク質を印付ける。しかしながら、モノユビキチン化に限定されるユビキチン化事象も一部存在し、その場合には1つのユビキチンのみがユビキチンリガーゼにより基質分子に付加される。モノユビキチン化されたタンパク質は、プロテアソームの分解標的とはならないが、そのかわりに例えばユビキチンを結合させることができるドメインを有する他のタンパク質との結合を介して、その細胞内の位置または機能を変化させる場合がある。さらに問題を複雑化しているのは、ユビキチン上の別のリシンがE3により標的とされ、鎖を形成し得るということである。最も共通したリシンは、ユビキチン鎖上のLys48である。これはプロテアソームにより認識されるポリユビキチンを生成するために使用されるリシンである。本明細書で使用される場合、「アルキル」という用語は、それ自体で、または別の置換基の一部として、別途明記されない限り、指定される炭素原子の数を有する直鎖または分岐鎖炭化水素ラジカルを意味する(すなわち、C1-8は、1~8個の炭素を意味する)。特定の数の炭素原子が存在しない場合、本明細書に提供されるアルキル基は、1~12個の炭素、1~8個の炭素、1~6個の炭素、または1~4個の炭素を有すると仮定される。アルキル基の例としては、メチル、エチル、n-プロピル、イソ-プロピル、n-ブチル、t-ブチル、イソ-ブチル、sec-ブチル、n-ペンチル、n-ヘキシル、n-ヘプチル、n-オクチルなどが挙げられる。アルキル基は、任意に、本明細書に提供されるように置換され得る。いくつかの実施形態では、アルキル基は、C1-6アルキルであり、いくつかの実施形態では、アルキル基は、C1-4アルキルである。 The term "ubiquitin ligase" refers to a family of proteins that facilitate the transfer of ubiquitin to a specific substrate protein, targeting the substrate protein for degradation. For example, E3 ubiquitin ligase proteins, alone or in combination with E2 ubiquitin conjugating enzymes, cause the attachment of ubiquitin to lysines on a target protein, which then targets the specific protein substrate for degradation by the proteasome. Thus, E3 ubiquitin ligases, alone or in complex with E2 ubiquitin conjugating enzymes, are involved in the transfer of ubiquitin to targeted proteins. Generally, ubiquitin ligases are involved in polyubiquitination, whereby a second ubiquitin is added to the first ubiquitin and a third ubiquitin is added to the second ubiquitin. Polyubiquitination marks a protein for proteasomal degradation. However, there are some ubiquitination events that are limited to monoubiquitination, where only one ubiquitin is added to the substrate molecule by the ubiquitin ligase. A monoubiquitinated protein is not targeted for degradation by the proteasome, but may instead change its cellular location or function, for example, through binding to other proteins that have domains to which ubiquitin can be attached. Further complicating the issue, additional lysines on the ubiquitin can be targeted by E3 to form chains. The most common lysine is Lys48 on the ubiquitin chain. This is the lysine used to generate polyubiquitin that is recognized by the proteasome. As used herein, the term "alkyl," by itself or as part of another substituent, means a straight or branched chain hydrocarbon radical having the number of carbon atoms designated, unless otherwise specified (i.e., C1-8 means 1 to 8 carbons). If no specific number of carbon atoms is present, the alkyl groups provided herein are assumed to have 1-12 carbons, 1-8 carbons, 1-6 carbons, or 1-4 carbons. Examples of alkyl groups include methyl, ethyl, n-propyl, iso-propyl, n-butyl, t-butyl, iso-butyl, sec-butyl, n-pentyl, n-hexyl, n-heptyl, n-octyl, and the like. The alkyl groups can be optionally substituted as provided herein. In some embodiments, the alkyl group is a C1-6 alkyl, and in some embodiments, the alkyl group is a C1-4 alkyl.
2016年8月5日に出願された米国特許出願第15/230,354号、および2014年7月11日に出願された米国特許出願第14/371,956号(米国特許出願公開第2014/0356322号として公開)、および2016年3月18日に出願された米国特許出願第15/074,820号(米国特許出願公開第2016/0272639号として公開)、および2016年2月24日に出願された国際特許出願第PCT/US2016/019328号(国際特許出願公開第WO2016/138114号として公開)、および2016年3月18日に出願された国際特許出願第PCT/US2016/023258号(国際特許出願公開第WO2016/149668号として公開)、2018年1月31日に出願された米国非仮特許出願第15/885,671号(米国特許出願公開第2018/0215731号として公開)、これらのすべては参照によりそれらの全体が本明細書に組み込まれる。さらに、本明細書に引用されるすべての参考文献は、参照によりそれらの全体が本明細書に組み込まれる。 U.S. Patent Application No. 15/230,354, filed August 5, 2016, U.S. Patent Application No. 14/371,956, filed July 11, 2014 (published as U.S. Patent Application Publication No. 2014/0356322), U.S. Patent Application No. 15/074,820, filed March 18, 2016 (published as U.S. Patent Application Publication No. 2016/0272639), and International Patent Application No. PCT/US2016/019, filed February 24, 2016 No. 328 (published as International Patent Application Publication No. WO2016/138114), and International Patent Application No. PCT/US2016/023258, filed March 18, 2016 (published as International Patent Application Publication No. WO2016/149668), and U.S. Nonprovisional Patent Application No. 15/885,671, filed January 31, 2018 (published as U.S. Patent Application Publication No. 2018/0215731), all of which are incorporated herein by reference in their entirety. Additionally, all references cited herein are incorporated herein by reference in their entirety.
「任意に置換された」という用語は、本明細書で定義される置換基と組み合わせて使用される場合、置換基が、本明細書に提供されるような1つ以上の好適な官能基または他の置換基で置換され得るが、必ずしもそうである必要はないことを意味する。例えば、置換基は、ハロ、シアノ、C1-6アルキル、C3-6シクロアルキル、C2-6アルケニル、C2-6アルキニル、ハロ(C1-6)アルキル、C1-6アルコキシ、ハロ(C1-6アルコキシ)、C1-6アルキルチオ、C1-6アルキルアミノ、NH2、NH(C1-6アルキル)、N(C1-6アルキル)2、NH(C1-6アルコキシ)、N(C1-6アルコキシ)2、-C(O)NHC1-6アルキル、-C(O)N(C1-6アルキル)2、-C(O)NH2、-C(O)C1-6アルキル、-C(O)2C1-6アルキル、-NHCO(C1-6アルキル)、-N(C1-6アルキル)CO(C1-6アルキル)、-S(O)C1-6アルキル、-S(O)2C1-6アルキル、オキソ、フェニル、ベンジル、ピリジニル、ピラゾリル、チアゾリル、イソチアゾリル、または他の5~6員ヘテロアリール基のうちの1つ以上で任意に置換され得る。いくつかの実施形態では、上記の任意の置換基の各々は、1個または2個の基で任意に置換されたそれら自身である。 The term "optionally substituted," when used in combination with a substituent defined herein, means that the substituent may, but need not, be substituted with one or more suitable functional groups or other substituents as provided herein. For example, the substituent may be selected from halo, cyano, C1-6 alkyl, C3-6 cycloalkyl, C2-6 alkenyl, C2-6 alkynyl, halo(C1-6)alkyl, C1-6 alkoxy, halo(C1-6 alkoxy), C1-6 alkylthio, C1-6 alkylamino, NH2, NH(C1-6 alkyl), N(C1-6 alkyl)2, NH(C1-6 alkoxy), N(C1-6 alkoxy)2, -C(O)NHC1-6 alkyl, -C(O) It may be optionally substituted with one or more of N(C1-6 alkyl)2, -C(O)NH2, -C(O)C1-6 alkyl, -C(O)2C1-6 alkyl, -NHCO(C1-6 alkyl), -N(C1-6 alkyl)CO(C1-6 alkyl), -S(O)C1-6 alkyl, -S(O)2C1-6 alkyl, oxo, phenyl, benzyl, pyridinyl, pyrazolyl, thiazolyl, isothiazolyl, or other 5-6 membered heteroaryl groups. In some embodiments, each of the above optional substituents is itself optionally substituted with one or two groups.
「シクロアルキル」という用語は、本明細書で使用される場合、C3-12環式アルキル基を指し、架橋およびスピロ環状(例えば、アダマンチン)を含む。シクロアルキル基の例としては、シクロプロピル、シクロブチル、シクロペンチル、シクロへプチル、シクロへキシル、シクロへプチル、シクロオクチル、ビシクロ[2.2.1]へプタニル、ビシクロ[3.1.1]へプタニル、ビシクロ[4.1.0]へプタニル、スピロ[3.3]へプタニル、およびスピロ[3.4]オクタニルが挙げられる。いくつかの実施形態では、シクロアルキル基は、C3-6シクロアルキルである。 The term "cycloalkyl" as used herein refers to a C3-12 cyclic alkyl group, including bridged and spirocyclic (e.g., adamantine). Examples of cycloalkyl groups include cyclopropyl, cyclobutyl, cyclopentyl, cycloheptyl, cyclohexyl, cycloheptyl, cyclooctyl, bicyclo[2.2.1]heptanyl, bicyclo[3.1.1]heptanyl, bicyclo[4.1.0]heptanyl, spiro[3.3]heptanyl, and spiro[3.4]octanyl. In some embodiments, the cycloalkyl group is a C3-6 cycloalkyl.
「アケニル」という用語は、本明細書で使用される場合、炭素原子の少なくとも2個がsp2ハイブリダイゼーションされ、それらの間に炭素-炭素二重結合を形成するC2-12アルキル基を指す。本明細書に提供されるアルケニル基は、2つ以上の炭素-炭素二重結合を含有し得るが、1つが好ましい。本明細書に提供されるアルケニル基のアルキル部分は、上記に提供されるように置換され得る。いくつかの実施形態では、アルケニル基は、C2-6アルケニルである。 The term "alkenyl," as used herein, refers to a C2-12 alkyl group in which at least two of the carbon atoms are sp2 hybridized to form a carbon-carbon double bond therebetween. The alkenyl groups provided herein can contain two or more carbon-carbon double bonds, although one is preferred. The alkyl portion of the alkenyl groups provided herein can be substituted as provided above. In some embodiments, the alkenyl group is a C2-6 alkenyl.
「アキニル」という用語は、本明細書で使用される場合、炭素原子の少なくとも2個がspハイブリダイゼーションされ、それらの間に炭素-炭素三重結合を形成するC2-12アルキル基を指す。本明細書に提供されるアルキニル基は、2つ以上の炭素-炭素三重結合を含有し得るが、1つが好ましい。本明細書に提供されるアルキニル基のアルキル部分は、上記に提供されるように置換され得る。いくつかの実施形態では、アルキニル基は、C2-6アルキニルである。 The term "alkynyl," as used herein, refers to a C2-12 alkyl group in which at least two of the carbon atoms are sp hybridized to form a carbon-carbon triple bond therebetween. The alkynyl groups provided herein can contain two or more carbon-carbon triple bonds, although one is preferred. The alkyl portion of the alkynyl groups provided herein can be substituted as provided above. In some embodiments, the alkynyl group is a C2-6 alkynyl.
「アルコキシ」、「アルキルアミノ」、および「アルキルチオ」という用語は、それらの従来の意味で使用され、酸素原子(「オキシ」)、アミノ基(「アミノ」)、またはチオ基を介して分子の残りに結合したアルキル基を指す。「アルキルアミノ」という用語は、モノ-ジ-アルキルアミノ基を含み、アルキル部分は同じであり得るか、または異なり得る。 The terms "alkoxy," "alkylamino," and "alkylthio" are used in their conventional sense to refer to an alkyl group attached to the remainder of the molecule via an oxygen atom ("oxy"), an amino group ("amino"), or a thio group. The term "alkylamino" includes mono- and di-alkylamino groups, where the alkyl portions may be the same or different.
単独での、または別の置換基の一部としての「ハロ」という用語は、別途明記されない限り、フッ素、塩素、臭素、またはヨウ素原子を意味するが、好ましくはフッ素または塩素を意味する。 The term "halo," by itself or as part of another substituent, means, unless otherwise stated, a fluorine, chlorine, bromine, or iodine atom, but preferably fluorine or chlorine.
「ハロ(C1-xアルキル)」という用語は、1~x個の炭素原子を有し、1個以上(例えば、1、2、3、4、5、または6個)のハロ基で置換されるアルキルを指す。例えば、この用語は、1個以上のハロ基で置換される1~6個の炭素原子を有するアルキル基を含む。ハロ(C1-C6アルキル)という用語の非限定的な例としては、フルオロメチル、ジフルオロメチル、トリフルオロメチル、クロロメチル、および2,2,2-トリフルオロエチルが挙げられる。 The term "halo(C1-x alkyl)" refers to an alkyl having 1 to x carbon atoms and substituted with one or more (e.g., 1, 2, 3, 4, 5, or 6) halo groups. For example, the term includes alkyl groups having 1 to 6 carbon atoms substituted with one or more halo groups. Non-limiting examples of the term halo(C1-C6 alkyl) include fluoromethyl, difluoromethyl, trifluoromethyl, chloromethyl, and 2,2,2-trifluoroethyl.
「ハロ(C1-xアルコキシ)」という用語は、1~x個の炭素原子を有し、1個以上(例えば、1、2、3、4、5、または6個)のハロ基で置換されるアルコキシ基を指す。例えば、この用語は、1個以上のハロ基で置換される1~6個の炭素原子を有するアルコキシ基を含む。ハロ(C1-C6アルキル)という用語の非限定的な例としては、フルオロメトキシ、ジフルオロメトキシ、トリフルオロメトキシ、クロロメトキシ、および2,2,2-トリフルオロエトキシが挙げられる。 The term "halo(C1-x alkoxy)" refers to an alkoxy group having 1 to x carbon atoms and substituted with one or more (e.g., 1, 2, 3, 4, 5, or 6) halo groups. For example, the term includes an alkoxy group having 1 to 6 carbon atoms substituted with one or more halo groups. Non-limiting examples of the term halo(C1-C6 alkyl) include fluoromethoxy, difluoromethoxy, trifluoromethoxy, chloromethoxy, and 2,2,2-trifluoroethoxy.
「ヘテロアルキル」という用語は、例えば、2~14個の炭素、例えば、2~10個の炭素を鎖中に有し、炭素の1個以上が、S、O、P、およびNから選択されるヘテロ原子によって置き換えられている、直鎖または分岐鎖アルキル基を指す。例示的なヘテロアルキルとしては、アルキルエーテル、二級および三級アルキルアミン、アルキルアミド、アルキルスルフィドなどが挙げられる。基は、末端基または架橋基であり得る。本明細書で使用される場合、架橋基の文脈で使用される場合の正常鎖への言及は、架橋基の2つの末端位置を結合する原子の直接鎖を指す。 The term "heteroalkyl" refers to a straight or branched chain alkyl group having, for example, 2 to 14 carbons, e.g., 2 to 10 carbons, in the chain, with one or more of the carbons replaced by a heteroatom selected from S, O, P, and N. Exemplary heteroalkyls include alkyl ethers, secondary and tertiary alkyl amines, alkyl amides, alkyl sulfides, and the like. The group may be a terminal group or a bridging group. As used herein, reference to a normal chain when used in the context of a bridging group refers to a direct chain of atoms connecting the two terminal positions of the bridging group.
「アリール」という用語は、本明細書で使用される場合、単一全炭素芳香族環または多重縮合全炭素環系を指し、環の少なくとも1つは芳香族である。例えば、ある特定の実施形態では、アリール基は、6~12個の炭素原子を有する。アリールには、フェニルラジカルが含まれる。アリールはまた、少なくとも1個の環が芳香族であり、他の環が芳香族であっても芳香族でなくてもよい、約9~12個の炭素原子を有する多重縮合環系(例えば、2、3、または4個の環を含む環系)を含む。このような多重縮合環系は、多重縮合環系の任意の炭素環部分上の1個以上(例えば、1、2、または3個)のオキソ基で任意に置換される。多重縮合環系の環は、価要件によって許容される場合、縮合結合、スピロ結合、および架橋結合を介して互いに連結することができる。多重縮合環系の結合点は、上記に定義されるように、環の芳香族または炭素環部分を含む環系の任意の位置にあり得ることを理解されたい。アリール基の非限定的な例としては、フェニル、インデニル、ナフチル、1、2、3、4-テトラヒドロナフチルなどが挙げられるが、これらに限定されない。 The term "aryl" as used herein refers to a single all-carbon aromatic ring or a multiple condensed all-carbon ring system, where at least one of the rings is aromatic. For example, in certain embodiments, an aryl group has 6-12 carbon atoms. Aryl includes phenyl radicals. Aryl also includes multiple condensed ring systems (e.g., ring systems containing 2, 3, or 4 rings) having about 9-12 carbon atoms, where at least one ring is aromatic and the other rings may or may not be aromatic. Such multiple condensed ring systems are optionally substituted with one or more (e.g., 1, 2, or 3) oxo groups on any carbocyclic portion of the multiple condensed ring system. The rings of the multiple condensed ring system can be linked to each other through fused bonds, spiro bonds, and bridge bonds, as permitted by valence requirements. It is understood that the point of attachment of the multiple condensed ring system can be at any position of the ring system, including the aromatic or carbocyclic portions of the ring, as defined above. Non-limiting examples of aryl groups include, but are not limited to, phenyl, indenyl, naphthyl, 1,2,3,4-tetrahydronaphthyl, and the like.
「ヘテロアリール」という用語は、本明細書で使用される場合、環内に炭素以外の少なくとも1個の原子を有する単一芳香族環を指し、原子は、酸素、窒素、および硫黄からなる群から選択され、「ヘテロアリール」はまた、少なくとも1個のこのような芳香族環を有する多重縮合環系も含み、多重縮合環系は、以下にさらに説明される。したがって、「ヘテロアリール」には、約1~6個の炭素原子、ならびに酸素、窒素、および硫黄からなる群から選択される約1~4個のヘテロ原子の単一芳香族環が含まれる。硫黄および窒素原子はまた、環が芳香族である限り、酸化形態で存在し得る。例示的なヘテロアリール環系としては、ピリジル、ピリミジニル、オキサゾリル、またはフリルが挙げられるが、これらに限定されない。「ヘテロアリール」はまた、多重縮合環系(例えば、2、3、または4個の環を含む環系)も含み、ヘテロアリール基は、上記に定義されるように、ヘテロアリール(例えば、1,8-ナフチリジニルなどのナフチリジニルを形成する)、複素環(例えば、1,2,3,4-テトラヒドロ-1,8-ナフチリジニルなどの1、2、3、4-テトラヒドロナフチリジニルを形成する)、炭素環(例えば、5,6,7,8-テトラヒドロキノリルを形成する)、およびアリール(例えば、インダゾリルを形成する)から選択される1つ以上の基と縮合して、多重縮合環系を形成する。したがって、ヘテロアリール(単一芳香族環または多重縮合環系)は、ヘテロアリール環内に約1~20個の炭素原子および約1~6個のヘテロ原子を有する。ヘテロアリール(単一芳香族環または多重縮合環系)はまた、ヘテロアリール環内に約5~12または約5~10員を有することもできる。多重縮合環系は、縮合環の炭素環または複素環部分上の1個以上(例えば、1、2、3、または4個)のオキソ基で任意に置換され得る。多重縮合環系の環は、価要件によって許容される場合、縮合結合、スピロ結合、および架橋結合を介して互いに連結することができる。多重縮合環系の個々の環は、互いに対して任意の順序で連結され得ることを理解されたい。また、多重縮合環系の結合点は(ヘテロアリールについて上記に定義されるように)、多重縮合環系のヘテロアリール、複素環、アリール、または炭素環部分を含む多重縮合環系の任意の位置にあり得ることも理解されたい。また、ヘテロアリールまたはヘテロアリール多重縮合環系の結合点は、炭素原子およびヘテロ原子(例えば、窒素)を含むヘテロアリールまたはヘテロアリール多重縮合環系の任意の好適な原子にあり得ることも理解されたい。例示的なヘテロアリールとしては、ピリジル、ピロリル、ピラジニル、ピリミジニル、ピリダジニル、ピラゾリル、チエニル、インドリル、イミダゾリル、オキサゾリル、イソキサゾリル、チアゾリル、フリル、オキサジアゾリル、チアジアゾリル、キノリル、イソキノリル、ベンゾチアゾリル、ベンゾキサゾリル、インダゾリル、キノキサリル、キナゾリル、5,6,7,8-テトラヒドロイソキノリニルベンゾフラニル、ベンズイミダゾリル、チアナフテニル、ピロロ[2,3-b]ピリジニル、キナゾリニル-4(3H)-オン、トリアゾリル、4,5,6,7-テトラヒドロ-1H-インダゾール、および3b,4,4a,5-テトラヒドロ-1H-シクロプロパ[3,4]シクロ-ペンタ[1,2-c]ピラゾールが挙げられるが、これらに限定されない。一実施形態では、「ヘテロアリール」という用語は、少なくとも1個のヘテロ原子を含有する単一芳香族環を指す。例えば、この用語は、1個以上のヘテロ原子を含む5員および6員の単環式芳香族環を含む。ヘテロアリールの非限定的な例としては、ピリジル、フリル、チアゾール、ピリミジン、オキサゾール、およびチアジアゾールが挙げられるが、これらに限定されない。 The term "heteroaryl" as used herein refers to a single aromatic ring having at least one atom other than carbon in the ring, the atom being selected from the group consisting of oxygen, nitrogen, and sulfur, and "heteroaryl" also includes multiple condensed ring systems having at least one such aromatic ring, which are further described below. Thus, "heteroaryl" includes a single aromatic ring of about 1 to 6 carbon atoms and about 1 to 4 heteroatoms selected from the group consisting of oxygen, nitrogen, and sulfur. The sulfur and nitrogen atoms may also be present in oxidized form, so long as the ring is aromatic. Exemplary heteroaryl ring systems include, but are not limited to, pyridyl, pyrimidinyl, oxazolyl, or furyl. "Heteroaryl" also includes multiple condensed ring systems (e.g., ring systems containing 2, 3, or 4 rings), where the heteroaryl group is fused to one or more groups selected from heteroaryl (e.g., to form naphthyridinyl, such as 1,8-naphthyridinyl), heterocycle (e.g., to form 1,2,3,4-tetrahydronaphthyridinyl, such as 1,2,3,4-tetrahydro-1,8-naphthyridinyl), carbocycle (e.g., to form 5,6,7,8-tetrahydroquinolyl), and aryl (e.g., to form indazolyl), as defined above, to form a multiple condensed ring system. Thus, a heteroaryl (single aromatic ring or multiple condensed ring system) has about 1-20 carbon atoms and about 1-6 heteroatoms in the heteroaryl ring. A heteroaryl (single aromatic ring or multiple condensed ring system) can also have about 5-12 or about 5-10 members in the heteroaryl ring. The multiple condensed ring system may be optionally substituted with one or more (e.g., 1, 2, 3, or 4) oxo groups on the carbocyclic or heterocyclic portion of the condensed ring. The rings of the multiple condensed ring system may be connected to each other through fused bonds, spiro bonds, and bridge bonds, if permitted by valence requirements. It is understood that the individual rings of the multiple condensed ring system may be connected to each other in any order. It is also understood that the point of attachment of the multiple condensed ring system (as defined above for heteroaryl) may be at any position of the multiple condensed ring system, including the heteroaryl, heterocyclic, aryl, or carbocyclic portion of the multiple condensed ring system. It is also understood that the point of attachment of the heteroaryl or heteroaryl multiple condensed ring system may be at any suitable atom of the heteroaryl or heteroaryl multiple condensed ring system, including carbon atoms and heteroatoms (e.g., nitrogen). Exemplary heteroaryls include, but are not limited to, pyridyl, pyrrolyl, pyrazinyl, pyrimidinyl, pyridazinyl, pyrazolyl, thienyl, indolyl, imidazolyl, oxazolyl, isoxazolyl, thiazolyl, furyl, oxadiazolyl, thiadiazolyl, quinolyl, isoquinolyl, benzothiazolyl, benzoxazolyl, indazolyl, quinoxalyl, quinazolyl, 5,6,7,8-tetrahydroisoquinolinylbenzofuranyl, benzimidazolyl, thianaphthenyl, pyrrolo[2,3-b]pyridinyl, quinazolinyl-4(3H)-one, triazolyl, 4,5,6,7-tetrahydro-1H-indazole, and 3b,4,4a,5-tetrahydro-1H-cyclopropa[3,4]cyclo-penta[1,2-c]pyrazole. In one embodiment, the term "heteroaryl" refers to a single aromatic ring containing at least one heteroatom. For example, the term includes 5- and 6-membered monocyclic aromatic rings containing one or more heteroatoms. Non-limiting examples of heteroaryls include, but are not limited to, pyridyl, furyl, thiazole, pyrimidine, oxazole, and thiadiazole.
「ヘテロシクリル」または「複素環」という用語は、本明細書で使用される場合、環内に炭素以外の少なくとも1個の原子を有する単一飽和または部分不飽和環を指し、原子は、酸素、窒素、および硫黄からなる群から選択され、この用語はまた、少なくとも1個のこのような飽和または部分不飽和環を有する多重縮合環系も含み、多重縮合環系は、以下にさらに説明される。したがって、この用語には、約1~6個の炭素原子、ならびに環中の酸素、窒素、および硫黄からなる群から選択される約1~3個のヘテロ原子の単一飽和または部分不飽和環(例えば、3、4、5、6、または7員環)が含まれる。環は、1個以上(例えば、1、2、または3個)のオキソ基で置換され得、硫黄および窒素原子もまた、それらの酸化形態で存在し得る。例示的な複素環としては、アゼチジニル、テトラヒドロフラニル、およびピペリジニルが挙げられるが、これらに限定されない。「複素環」という用語はまた、多重縮合環系(例えば、2、3、または4個の環を含む環系)も含み、単一複素環は(上記に定義されるように)、複素環(例えば、1,8-デカヒドロナプチリジニルを形成する)、炭素環(例えば、デカヒドロキノリルを形成する)、およびアリールから選択される1つ以上の基と縮合して、多重縮合環系を形成することができる。したがって、複素環(単一飽和もしくは単一部分不飽和環、または多重縮合環系)は、複素環内に約2~20個の炭素原子および1~6個のヘテロ原子を有する。このような多重縮合環系は、多重縮合環の炭素環または複素環部分上の1個以上(例えば、1、2、3、または4個)のオキソ基で任意に置換され得る。多重縮合環系の環は、価要件によって許容される場合、縮合結合、スピロ結合、および架橋結合を介して互いに連結することができる。多重縮合環系の個々の環は、互いに対して任意の順序で連結され得ることを理解されたい。したがって、複素環(単一飽和もしくは単一部分不飽和環、または多重縮合環系)は、複素環系内に約1~6個のヘテロ原子を含む約3~20個の原子を有する。また、多重縮合環系の結合点は(複素環について上記に定義されるように)、環の複素環、アリール、および炭素環部分を含む多重縮合環系の任意の場所にあり得ることも理解されたい。また、複素環または複素環多重縮合環系の結合点は、炭素原子およびヘテロ原子(例えば、窒素)を含む複素環または複素環多重縮合環系の任意の好適な原子にあり得ることも理解されたい。一実施形態では、複素環という用語は、C2-20複素環を含む。一実施形態では、複素環という用語は、C2-7複素環を含む。一実施形態では、複素環という用語は、C2-5複素環を含む。一実施形態では、複素環という用語は、C2-4複素環を含む。例示的な複素環としては、アジリジニル、アゼチジニル、ピロリジニル、ピペリジニル、ホモピペリジニル、モルホリニル、チオモルホリニル、ピペラジニル、テトラヒドロフラニル、ジヒドロオキサゾリル、テトラヒドロピラニル、テトラヒドロチオピラニル、1,2,3,4-テトラヒドロキノリル、ベンゾキサジニル、ジヒドロオキサゾリル、クロマニル、1,2-ジヒドロピリジニル、2,3-ジヒドロベンゾフラニル、1,3-ベンゾジオキソリル、1,4-ベンゾジオキサニル、スピロ[シクロプロパン-1,1’-イソインドリニル]-3’-オン、イソインドリニル-1-オン、2-オキサ-6-アザスピロ[3.3]ヘプタニル、イミダゾリジン-2-オンN-メチルピペリジン、イミダゾリジン、ピラゾリジン、ブチロラクタム、バレロラクタム、イミダゾリジノン、ヒダントイン、ジオキソラン、フタルイミド、1,4-ジオキサン、チオモルホリン、チオモルホリンS-オキシド、チオモルホリンS,S-オキシド、ピラン、3-ピロリン、チオピラン、ピロン、テトラヒドロチオフェン、キヌクリジン、トロパン、2-アザスピロ[3.3]ヘプタン、(1R,5S)-3-アザビシクロ[3.2.1]オクタン、(1s,4s)-2-アザビシクロ[2.2.2]オクタン、(1R,4R)-2-オキサ-5-アザビシクロ[2.2.2]オクタンおよび、ピロリジン-2-オンが挙げられるが、これらに限定されない。一実施形態では、「複素環」という用語は、少なくとも1つのヘテロ原子を有する単環式、飽和または部分不飽和3~8員環を指す。例えば、この用語は、少なくとも1つのヘテロ原子を有する単環式、飽和または部分不飽和4、5、6、または7員環を含む。複素環の非限定的な例としては、アジリジン、アゼチジン、ピロリジン、ピペリジン、ピペリジン、ピペラジン、オキシラン、モルホリン、およびチオモルホリンが挙げられる。本明細書で使用される場合、「9員または10員複素環」という用語は、少なくとも1つのヘテロ原子を有する部分不飽和または芳香族縮合二環式環系を指す。例えば、9員または10員複素環という用語は、1つ以上のヘテロ原子を含有する5員または6員飽和、部分不飽和、または芳香族環に縮合されたベンゾ環を有する二環式環系を含む。 The term "heterocyclyl" or "heterocycle" as used herein refers to a single saturated or partially unsaturated ring having at least one atom other than carbon in the ring, the atom being selected from the group consisting of oxygen, nitrogen, and sulfur, and the term also includes multiple condensed ring systems having at least one such saturated or partially unsaturated ring, which are further described below. Thus, the term includes a single saturated or partially unsaturated ring (e.g., a 3-, 4-, 5-, 6-, or 7-membered ring) of about 1 to 6 carbon atoms and about 1 to 3 heteroatoms selected from the group consisting of oxygen, nitrogen, and sulfur in the ring. The ring may be substituted with one or more (e.g., 1, 2, or 3) oxo groups, and the sulfur and nitrogen atoms may also be present in their oxidized forms. Exemplary heterocycles include, but are not limited to, azetidinyl, tetrahydrofuranyl, and piperidinyl. The term "heterocycle" also includes multiple condensed ring systems (e.g., ring systems containing 2, 3, or 4 rings), where a single heterocycle (as defined above) can be fused with one or more groups selected from heterocycle (e.g., to form 1,8-decahydronapthyridinyl), carbocycle (e.g., to form decahydroquinolyl), and aryl to form a multiple condensed ring system. Thus, a heterocycle (a single saturated or partially unsaturated ring, or a multiple condensed ring system) has from about 2 to 20 carbon atoms and from 1 to 6 heteroatoms in the heterocycle. Such multiple condensed ring systems can be optionally substituted with one or more (e.g., 1, 2, 3, or 4) oxo groups on the carbocycle or heterocycle portions of the multiple condensed ring. The rings of a multiple condensed ring system can be connected to each other through fused bonds, spiro bonds, and bridge bonds, where permitted by valency requirements. It is understood that the individual rings of a multiple condensed ring system can be connected to each other in any order. Thus, a heterocyclic ring (single saturated or single partially unsaturated ring, or multiple condensed ring system) has from about 3 to 20 atoms, including from about 1 to 6 heteroatoms in the heterocyclic ring system. It is also understood that the point of attachment of the multiple condensed ring system (as defined above for heterocyclic rings) can be anywhere in the multiple condensed ring system, including the heterocyclic, aryl, and carbocyclic portions of the ring. It is also understood that the point of attachment of a heterocyclic ring or heterocyclic multiple condensed ring system can be at any suitable atom of the heterocyclic ring or heterocyclic multiple condensed ring system, including carbon atoms and heteroatoms (e.g., nitrogen). In one embodiment, the term heterocyclic includes a C2-20 heterocyclic ring. In one embodiment, the term heterocyclic includes a C2-7 heterocyclic ring. In one embodiment, the term heterocyclic includes a C2-5 heterocyclic ring. In one embodiment, the term heterocyclic includes a C2-4 heterocyclic ring. Exemplary heterocycles include aziridinyl, azetidinyl, pyrrolidinyl, piperidinyl, homopiperidinyl, morpholinyl, thiomorpholinyl, piperazinyl, tetrahydrofuranyl, dihydrooxazolyl, tetrahydropyranyl, tetrahydrothiopyranyl, 1,2,3,4-tetrahydroquinolyl, benzoxazinyl, dihydrooxazolyl, chromanyl, 1,2-dihydropyridinyl, 2,3-dihydrobenzofuranyl, 1,3-benzodioxolyl, 1,4-benzodioxanyl, spiro[cyclopropane-1,1'-isoindolinyl]-3'-one, isoindolinyl-1-one, 2-oxa-6-azaspiro[3.3]heptanyl, imidazolidin-2-one, These include, but are not limited to, N-methylpiperidine, imidazolidine, pyrazolidine, butyrolactam, valerolactam, imidazolidinone, hydantoin, dioxolane, phthalimide, 1,4-dioxane, thiomorpholine, thiomorpholine S-oxide, thiomorpholine S,S-oxide, pyran, 3-pyrroline, thiopyran, pyrone, tetrahydrothiophene, quinuclidine, tropane, 2-azaspiro[3.3]heptane, (1R,5S)-3-azabicyclo[3.2.1]octane, (1s,4s)-2-azabicyclo[2.2.2]octane, (1R,4R)-2-oxa-5-azabicyclo[2.2.2]octane, and pyrrolidin-2-one. In one embodiment, the term "heterocycle" refers to a monocyclic, saturated or partially unsaturated 3-8 membered ring having at least one heteroatom. For example, the term includes monocyclic, saturated or partially unsaturated 4-, 5-, 6-, or 7-membered rings having at least one heteroatom. Non-limiting examples of heterocycles include aziridine, azetidine, pyrrolidine, piperidine, piperazine, oxirane, morpholine, and thiomorpholine. As used herein, the term "9- or 10-membered heterocycle" refers to a partially unsaturated or aromatic fused bicyclic ring system having at least one heteroatom. For example, the term 9- or 10-membered heterocycle includes bicyclic ring systems having a benzo ring fused to a 5- or 6-membered saturated, partially unsaturated, or aromatic ring containing one or more heteroatoms.
本明細書で使用される場合、「ヘテロ原子」という用語は、酸素(O)、窒素(N)、硫黄(S)、およびケイ素(Si)を含むことを意味する。窒素および硫黄は、可能な場合、酸化形態であり得る。 As used herein, the term "heteroatom" is meant to include oxygen (O), nitrogen (N), sulfur (S), and silicon (Si). Nitrogen and sulfur may be in oxidized form, where possible.
本明細書で使用される場合、「キラル」という用語は、鏡像パートナーに重ねることができない特性を有する分子を指し、一方、「アキラル」という用語は、それらの鏡像パートナーに重ねることができる分子を指す。 As used herein, the term "chiral" refers to molecules that have the property of not being superimposable on their mirror image partners, while the term "achiral" refers to molecules that can be superimposed on their mirror image partners.
本明細書で使用される場合、「立体異性体」という用語は、同一の化学構造を有するが、空間における原子または基の配置の点で異なる化合物を指す。本明細書で使用される場合、交差線「
本明細書で使用される場合、化学構造中の結合と交差する波線「
「ジアステレオマー」とは、2つ以上のキラリティーの中心を有し、その分子が互いの鏡像ではない立体異性体を指す。ジアステレオマーは、異なる物理的特性、例えば、融点、沸点、スペクトル特性、および反応性を有する。ジアステレオマーの混合物は、電気泳動およびクロマトグラフィーなどの高分解能分析手法下で分離することができる。 "Diastereomer" refers to a stereoisomer with two or more centers of chirality and whose molecules are not mirror images of one another. Diastereomers have different physical properties, e.g., melting points, boiling points, spectral properties, and reactivities. Mixtures of diastereomers can separate under high resolution analytical techniques such as electrophoresis and chromatography.
「鏡像異性体」とは、互いに重ねることができない鏡像である化合物の2つの立体異性体を指す。 "Enantiomers" refers to two stereoisomers of a compound that are non-superimposable mirror images of one another.
本明細書に使用される立体化学的定義および慣習は、概して、S.P.Parker,Ed.,McGraw-Hill Dictionary of Chemical Terms(1984)McGraw-Hill Book Company,New York、およびEliel,E.and Wilen,S.,“Stereochemistry of Organic Compounds”,John Wiley & Sons,Inc.,New York,1994に従う。本発明の化合物は、非対称またはキラル中心を含有することができ、したがって、異なる立体異性体形態で存在する。ジアステレオマー、鏡像異性体、およびアトロプ異性体、ならびにラセミ混合物などのそれらの混合物を含むが、これらに限定されない、本発明の化合物のすべての立体異性体形態が、本発明の一部を形成することが意図される。多くの有機化合物は光学活性形態で存在し、すなわち、それらは、平面偏光の平面を回転させる能力を有する。光学活性化合物の説明において、接頭辞DおよびL、またはRおよびSは、キラル中心の周りの分子の絶対配置を示すために使用される。接頭辞dおよびlまたは(+)および(-)は、化合物による平面偏光の回転の印を示すために用いられ、(-)または1は、化合物が左旋性であることを意味する。(+)またはdの接頭辞を有する化合物は、右旋性である。所与の化学構造について、これらの立体異性体は、それらが互いの鏡像であるということ以外、同一である。特定の立体異性体はまた、鏡像異性体と称することもでき、このような異性体の混合物は、多くの場合、鏡像異性混合物と呼ばれる。鏡像異性体の50:50の混合物は、ラセミ混合物またはラセミ体と称され、これは、化学反応またはプロセスにおいて立体選択または立体特異性が存在しなかった場合に生じ得る。「ラセミ混合物」および「ラセミ体」という用語は、光学活性を欠く2つの鏡像異性種の等モル混合物を指す。 Stereochemical definitions and conventions used herein generally follow S. P. Parker, Ed., McGraw-Hill Dictionary of Chemical Terms (1984) McGraw-Hill Book Company, New York, and Eliel, E. and Wilen, S., "Stereochemistry of Organic Compounds", John Wiley & Sons, Inc., New York, 1994. The compounds of the invention may contain asymmetric or chiral centers and therefore exist in different stereoisomeric forms. All stereoisomeric forms of the compounds of the present invention, including, but not limited to, diastereomers, enantiomers, and atropisomers, as well as mixtures thereof, such as racemic mixtures, are intended to form part of the present invention. Many organic compounds exist in optically active forms, i.e., they have the ability to rotate the plane of plane-polarized light. In describing optically active compounds, the prefixes D and L, or R and S, are used to indicate the absolute configuration of the molecule about a chiral center. The prefixes d and l, or (+) and (-), are used to indicate the sign of rotation of plane-polarized light by the compound, with (-) or 1 meaning that the compound is levorotatory. Compounds prefixed with (+) or d are dextrorotatory. For a given chemical structure, these stereoisomers are identical except that they are mirror images of one another. A specific stereoisomer can also be referred to as an enantiomer, and a mixture of such isomers is often called an enantiomeric mixture. A 50:50 mixture of enantiomers is called a racemic mixture or racemate, which may occur where there has been no stereoselection or stereospecificity in a chemical reaction or process. The terms "racemic mixture" and "racemate" refer to an equimolar mixture of two enantiomeric species, devoid of optical activity.
本明細書の化合物式中の結合が非立体化学的様式(例えば、平坦)で描画される場合、結合が結合している原子は、すべての立体化学的可能性を含む。本明細書の化合物式中の結合が、規定の立体化学的様式(例えば、太字、太字くさび、破線または破線くさび)で描画される場合、別途記載がない限り、立体化学的結合が結合する原子が、描写される絶対立体異性体中で濃縮されることを理解されたい。一実施形態では、化合物は、示される絶対立体異性体の少なくとも51%であり得る。別の実施形態では、化合物は、示される絶対立体異性体の少なくとも80%であり得る。別の実施形態では、化合物は、示される絶対立体異性体の少なくとも90%であり得る。別の実施形態では、化合物は、示される絶対立体異性体の少なくとも95%であり得る。別の実施形態では、化合物は、示される絶対立体異性体の少なくとも97%であり得る。別の実施形態では、化合物は、示される絶対立体異性体の少なくとも98%であり得る。別の実施形態では、化合物は、示される絶対立体異性体の少なくとも99%であり得る。 When bonds in the compound formulae herein are drawn in a non-stereochemical manner (e.g., flat), the atoms to which the bonds are attached include all stereochemical possibilities. When bonds in the compound formulae herein are drawn in a prescribed stereochemical manner (e.g., bold, bold wedge, dashed line, or dashed wedge), it is understood that the atoms to which the stereochemical bonds are attached are enriched in the depicted absolute stereoisomer, unless otherwise noted. In one embodiment, the compound may be at least 51% of the absolute stereoisomer shown. In another embodiment, the compound may be at least 80% of the absolute stereoisomer shown. In another embodiment, the compound may be at least 90% of the absolute stereoisomer shown. In another embodiment, the compound may be at least 95% of the absolute stereoisomer shown. In another embodiment, the compound may be at least 97% of the absolute stereoisomer shown. In another embodiment, the compound may be at least 98% of the absolute stereoisomer shown. In another embodiment, the compound may be at least 99% of the absolute stereoisomer shown.
本明細書で使用される場合、「互変異性体」または「互変異性形態」という用語は、低エネルギー障壁を介して相互変換可能である、異なるエネルギーの構造異性体を指す。例えば、プロトン互変異性体(プロトトロピック互変異性体としても既知)は、ケト-エノールおよびイミン-エナミン異性化などのプロトンの移行を介した相互変換を含む。原子価互変異性体は、結合電子のうちのいくつかの再編成による相互変換を含む。 As used herein, the term "tautomer" or "tautomeric form" refers to structural isomers of different energies that are interconvertible via a low energy barrier. For example, proton tautomers (also known as prototropic tautomers) include interconversions via migration of a proton, such as keto-enol and imine-enamine isomerizations. Valence tautomers include interconversions by rearrangement of some of the bonding electrons.
本明細書で使用される場合、「溶媒和物」という用語は、1つ以上の溶媒分子と、本発明の化合物との会合または複合体を指す。溶媒和物を形成する溶媒の例としては、水、イソプロパノール、エタノール、メタノール、DMSO、酢酸エチル、酢酸、およびエタノールアミンが挙げられるが、これらに限定されない。「水和物」という用語は、溶媒分子が水である複合体を指す。 As used herein, the term "solvate" refers to an association or complex of one or more solvent molecules with a compound of the invention. Examples of solvents that form solvates include, but are not limited to, water, isopropanol, ethanol, methanol, DMSO, ethyl acetate, acetic acid, and ethanolamine. The term "hydrate" refers to a complex where the solvent molecule is water.
本明細書で使用される場合、「保護基」という用語は、化合物上の特定の官能基を遮断または保護するために一般的に用いられる置換基を指す。例えば、「アミノ保護基」は、化合物中のアミノ官能基を遮断または保護する、アミノ基に結合される置換基である。好適なアミノ保護基としては、アセチル、トリフルオロアセチル、t-ブトキシカルボニル(BOC)、ベンジルオキシカルボニル(CBZ)、および9-フルオレニルメチレンオキシカルボニル(Fmoc)が挙げられる。同様に、「ヒドロキシ保護基」とは、ヒドロキシ官能基を遮断または保護する、ヒドロキシ基の置換基を指す。好適な保護基としては、アセチルおよびシリルが挙げられる。「カルボキシ保護基」は、カルボキシ官能基を遮断または保護する、カルボキシ基の置換基を指す。一般的なカルボキシ保護基としては、フェニルスルホニルエチル、シアノエチル、2-(トリメチルシリル)エチル、2-(トリメチルシリル)エトキシメチル、2-(p-トルエンスルホニル)エチル、2-(p-ニトロフェニルスルフェニル)エチル、2-(ジフェニルホスフィノ)-エチル、ニトロエチルなどが挙げられる。保護基およびそれらの使用の概要については、P.G.M.WutsおよびT.W.Greene,Greene’s Protective Groups in Organic Synthesis 4th edition,Wiley-Interscience,New York,2006を参照されたい。 As used herein, the term "protecting group" refers to a substituent that is commonly used to block or protect a particular functional group on a compound. For example, an "amino protecting group" is a substituent attached to an amino group that blocks or protects the amino functionality in the compound. Suitable amino protecting groups include acetyl, trifluoroacetyl, t-butoxycarbonyl (BOC), benzyloxycarbonyl (CBZ), and 9-fluorenylmethyleneoxycarbonyl (Fmoc). Similarly, a "hydroxy protecting group" refers to a substituent of a hydroxy group that blocks or protects the hydroxy functionality. Suitable protecting groups include acetyl and silyl. A "carboxy protecting group" refers to a substituent of a carboxy group that blocks or protects the carboxy functionality. Common carboxy protecting groups include phenylsulfonylethyl, cyanoethyl, 2-(trimethylsilyl)ethyl, 2-(trimethylsilyl)ethoxymethyl, 2-(p-toluenesulfonyl)ethyl, 2-(p-nitrophenylsulfenyl)ethyl, 2-(diphenylphosphino)-ethyl, nitroethyl, and the like. For a general review of protecting groups and their uses, see P. G. M. Wuts and T. W. Greene, Greene's Protective Groups in Organic Synthesis 4th edition, Wiley-Interscience, New York, 2006.
本明細書で使用される場合、「薬学的に許容される塩」という用語は、本明細書に記載される化合物に見出される特定の置換基に応じて、比較的非毒性の酸または塩基で調製される活性化合物の塩を含むことを意味する。本発明の化合物が比較的酸性の官能基を含有する場合、塩基付加塩は、そのような化合物の中性形態を、十分な量の所望の塩基と、そのままで、または好適な不活性溶媒中のいずれかで接触させることによって得ることができる。薬学的に許容される無機塩基に由来する塩の例としては、アルミニウム、アンモニウム、カルシウム、銅、第二鉄、第一鉄、リチウム、マグネシウム、第二マンガン、第一マンガン、カリウム、ナトリウム、亜鉛などが挙げられる。薬学的に許容される有機塩基に由来する塩としては、置換アミン、環状アミン、天然型アミンなどを含む、第一級、第二級、および第三級アミンの塩、例えば、アルギニン、ベタイン、カフェイン、コリン、N,N’-ジベンジルエチレンジアミン、ジエチルアミン、2-ジエチルアミノエタノール、2-ジメチルアミノエタノール、エタノールアミン、エチレンジアミン、N-エチルモルホリン、N-エチルピペリジン、グルカミン、グルコサミン、ヒスチジン、ヒドラバミン、イソプロピルアミン、リジン、メチルグルカミン、モルホリン、ピペラジン、ピペリジン、ポリアミン樹脂、プロカイン、プリン、テオブロミン、トリエチルアミン、トリメチルアミン、トリプロピルアミン、トロメタミンなどが挙げられる。本発明の化合物が比較的塩基性の官能基を含有する場合、酸付加塩は、そのような化合物の中性形態を、十分な量の所望の酸と、そのままで、または好適な不活性溶媒中のいずれかで接触させることによって得ることができる。薬学的に許容される酸付加塩の例としては、塩酸、臭化水素酸、硝酸、炭酸、一水素炭酸、リン酸、一水素リン酸、二水素リン酸、硫酸、一水素硫酸、ヨウ化水素酸、またはリン酸などの無機酸に由来する塩、ならびに酢酸、プロピオン酸、イソ酪酸、マロン酸、安息香酸、コハク酸、スベリン酸、フマル酸、マンデル酸、フタル酸、ベンゼンスルホン酸、p-トリルスルホン酸、クエン酸、酒石酸、メタンスルホン酸などの比較的非毒性の有機酸に由来する塩が挙げられる。また、アルギニン酸などのアミノ酸の塩、およびグルコロン酸またはガラクツロン酸などの有機酸の塩も含まれる(例えば、Berge,S.M.,et al.,「Pharmaceutical Salts」,Journal of Pharmaceutical Science,1977,66,1-19参照)。本発明のある特定の具体的な化合物は、化合物を塩基または酸付加塩のいずれかに変換することを可能にする塩基性官能基および酸性官能基の両方を含有する。 As used herein, the term "pharmaceutical acceptable salts" is meant to include salts of active compounds prepared with relatively non-toxic acids or bases, depending on the particular substituents found on the compounds described herein. When the compounds of the present invention contain relatively acidic functional groups, base addition salts can be obtained by contacting the neutral form of such compounds with a sufficient amount of the desired base, either neat or in a suitable inert solvent. Examples of salts derived from pharmaceutical acceptable inorganic bases include aluminum, ammonium, calcium, copper, ferric, ferrous, lithium, magnesium, manganic, manganous, potassium, sodium, zinc, and the like. Salts derived from pharma- ceutically acceptable organic bases include salts of primary, secondary, and tertiary amines, including substituted amines, cyclic amines, naturally occurring amines, and the like, such as arginine, betaine, caffeine, choline, N,N'-dibenzylethylenediamine, diethylamine, 2-diethylaminoethanol, 2-dimethylaminoethanol, ethanolamine, ethylenediamine, N-ethylmorpholine, N-ethylpiperidine, glucamine, glucosamine, histidine, hydrabamine, isopropylamine, lysine, methylglucamine, morpholine, piperazine, piperidine, polyamine resins, procaine, purine, theobromine, triethylamine, trimethylamine, tripropylamine, tromethamine, and the like. When a compound of the invention contains a relatively basic functional group, an acid addition salt can be obtained by contacting the neutral form of such a compound with a sufficient amount of the desired acid, either neat or in a suitable inert solvent. Examples of pharma-ceutically acceptable acid addition salts include salts derived from inorganic acids such as hydrochloric, hydrobromic, nitric, carbonic, monohydrogencarbonic, phosphoric, monohydrogenphosphate, dihydrogenphosphate, sulfuric, monohydrogensulfuric, hydroiodic, or phosphoric acid, as well as salts derived from relatively non-toxic organic acids such as acetic, propionic, isobutyric, malonic, benzoic, succinic, suberic, fumaric, mandelic, phthalic, benzenesulfonic, p-tolylsulfonic, citric, tartaric, methanesulfonic, etc. Also included are salts of amino acids such as arginic acid, and salts of organic acids such as glucoronic or galacturonic acid (see, e.g., Berge, S.M., et al., "Pharmaceutical Salts", Journal of Pharmaceutical Science, 1977, 66, 1-19). Certain specific compounds of the present invention contain both basic and acidic functionalities that allow the compounds to be converted into either base or acid addition salts.
化合物の中性形態は、塩を塩基または酸と接触させ、従来の方法で親化合物を単離することによって再生することができる。化合物の親形態は、ある特定の物理的特性、例えば、極性溶媒中の溶解性において、様々な塩形態と異なるが、そうでなければ、塩は、本発明の目的のための化合物の親形態と同等である。 The neutral forms of the compounds can be regenerated by contacting the salt with a base or acid and isolating the parent compound in the conventional manner. The parent form of the compound differs from the various salt forms in certain physical properties, e.g., solubility in polar solvents, but otherwise the salts are equivalent to the parent form of the compound for purposes of this invention.
塩形態に加えて、本発明は、プロドラッグ形態である化合物を提供する。本明細書で使用される場合、「プロドラッグ」という用語は、本発明の化合物を提供するために生理学的条件下で化学変化を容易に受ける化合物を指す。さらに、プロドラッグは、エクスビボ環境において化学的または生化学的方法によって本発明の化合物に変換することができる。例えば、プロドラッグは、好適な酵素または化学試薬とともに経皮パッチリザーバに配置される場合、本発明の化合物にゆっくりと変換することができる。 In addition to salt forms, the present invention provides compounds that are in prodrug form. As used herein, the term "prodrug" refers to a compound that readily undergoes chemical changes under physiological conditions to provide a compound of the present invention. Additionally, prodrugs can be converted to the compounds of the present invention by chemical or biochemical methods in an ex vivo environment. For example, prodrugs can be slowly converted to the compounds of the present invention when placed in a transdermal patch reservoir with a suitable enzyme or chemical reagent.
本発明のプロドラッグには、アミノ酸残基、または2個以上(例えば、2個、3個、または4個)のアミノ酸残基のポリペプチド鎖が、アミドまたはエステル結合によって、本発明の化合物の遊離アミノ、ヒドロキシ、またはカルボン酸基に共有結合している化合物が含まれる。アミノ酸残基には、3文字の記号によって一般的に指定される20個の天然型アミノ酸が含まれるが、これらに限定されず、また、ホスホセリン、ホスホトレオニン、ホスホチロシン、4-ヒドロキシプロリン、ヒドロキシリジン、デモシン、イソデモシン、ガンマ-カルボキシグルタミン酸、ヒプリン酸、オクタヒドロインドール-2-カルボン酸、スタチン、1,2,3,4-テトラヒドロイソキノリン-3-カルボン酸、ペニシルアミン、オルニチン、3-メチルヒスチジン、ノルバリン、ベータ-アラニン、ガンマ-アミノブチリン酸、シトルリン、ホモシステイン、ホモセリン、メチルアラニン、パラ-ベンゾイルフェニルアラニン、フェニルグリシン、プロパルギルグリシン、サルコシン、メチオニンスルホン、およびtert-ブチルグリシンも含まれる。 Prodrugs of the present invention include compounds in which an amino acid residue, or a polypeptide chain of two or more (e.g., two, three, or four) amino acid residues, is covalently bonded to a free amino, hydroxy, or carboxylic acid group of a compound of the present invention via an amide or ester bond. Amino acid residues include, but are not limited to, the 20 naturally occurring amino acids commonly designated by their three letter symbols, and also include phosphoserine, phosphothreonine, phosphotyrosine, 4-hydroxyproline, hydroxylysine, decomine, isodecomine, gamma-carboxyglutamic acid, hippuric acid, octahydroindole-2-carboxylic acid, statin, 1,2,3,4-tetrahydroisoquinoline-3-carboxylic acid, penicillamine, ornithine, 3-methylhistidine, norvaline, beta-alanine, gamma-aminobutyric acid, citrulline, homocysteine, homoserine, methylalanine, para-benzoylphenylalanine, phenylglycine, propargylglycine, sarcosine, methionine sulfone, and tert-butylglycine.
追加の種類のプロドラッグもまた包含される。例えば、本発明の化合物の遊離カルボキシル基は、アミドまたはアルキルエステルとして誘導体化することができる。別の例として、遊離ヒドロキシ基を含む本発明の化合物は、Fleisher,D.et al.,(1996)Improved oral drug delivery:solubility limitations overcome by the use of prodrugs Advanced Drug Delivery Reviews,19:115に概説されるように、ヒドロキシ基を、これらに限定されないが、リン酸エステル、ヘミスクシナート、ジメチルアミノアセテート、またはホスホリルオキシメチルオキシカルボニル基などの基に変換することによって、プロドラッグとして誘導体化することができる。ヒドロキシおよびアミノ基のカルバメートプロドラッグには、カルボネートプロドラッグとして、ヒドロキシ基のスルホン酸エステルおよび硫酸エステルも含まれる。(アシルオキシ)メチルおよび(アシルオキシ)エチルエーテルとしてのヒドロキシ基の誘導体化も包含され、アシル基は、エーテル、アミン、およびカルボン酸官能基を含むがこれらに限定されない基で任意に置換されたアルキルエステルであり得るか、またはアシル基は、上記に記載されたアミノ酸エステルである。この種類のプロドラッグは、J.Med.Chem.,(1996),39:10に記載されている。より具体的な例としては、アルコール基の水素原子を、(C1-6)アルカノイルオキシメチル、1-((C1-6)アルカノイルオキシ)エチル、1-メチル-1-((C1-6)アルカノイルオキシ)エチル、(C1-6)アルコキシカルボニルオキシメチル、N-(C1-6)アルコキシカルボニルアミノメチル、スクシノイル、(C1-6)アルカノイル、アルファ-アミノ(C1-4)アルカノイル、アリルアシル、およびアルファ-アミノアシル、またはアルファ-アミノアシル-アルファ-アミノアシルなどの基で置き換えることを含み、各アルファ-アミノアシル基は、独立して、天然型L-アミノ酸、P(O)(OH)2、-P(O)(O(C1-6)アルキル)2、またはグリコシル(炭水化物のヘミアセタール形態のヒドロキシル基の除去に起因するラジカル)から選択される。 Additional types of prodrugs are also included. For example, free carboxyl groups of the compounds of the invention can be derivatized as amides or alkyl esters. As another example, compounds of the invention containing free hydroxyl groups can be derivatized as prodrugs by converting the hydroxyl group to groups such as, but not limited to, phosphate esters, hemisuccinates, dimethylaminoacetates, or phosphoryloxymethyloxycarbonyl groups, as reviewed in Fleisher, D. et al., (1996) Improved oral drug delivery: solubility limitations overcome by the use of prodrugs Advanced Drug Delivery Reviews, 19:115. Carbamate prodrugs of hydroxy and amino groups also include sulfonate and sulfate esters of hydroxy groups, as well as carbonate prodrugs.Derivatization of hydroxy groups as (acyloxy)methyl and (acyloxy)ethyl ethers is also included, and acyl groups can be alkyl esters, optionally substituted with groups including but not limited to ether, amine, and carboxylic acid functional groups, or acyl groups are amino acid esters as described above.This type of prodrug is described in J. Med. Chem., (1996), 39:10. More specific examples include replacing the hydrogen atom of the alcohol group with groups such as (C1-6)alkanoyloxymethyl, 1-((C1-6)alkanoyloxy)ethyl, 1-methyl-1-((C1-6)alkanoyloxy)ethyl, (C1-6)alkoxycarbonyloxymethyl, N-(C1-6)alkoxycarbonylaminomethyl, succinoyl, (C1-6)alkanoyl, alpha-amino(C1-4)alkanoyl, aryl acyl, and alpha-aminoacyl, or alpha-aminoacyl-alpha-aminoacyl, each alpha-aminoacyl group being independently selected from naturally occurring L-amino acids, P(O)(OH)2, -P(O)(O(C1-6)alkyl)2, or glycosyl (the radical resulting from removal of the hydroxyl group of the hemiacetal form of a carbohydrate).
プロドラッグ誘導体の追加の例については、例えば、a)Design of Prodrugs、H.Bundgaard編(Elsevier,1985)、およびMethods in Enzymology,Vol.42,p.309-396、K.Widder,et al.編(Academic Press,1985)、b)A Textbook of Drug Design and Development、Krogsgaard-LarsenおよびH.Bundgaard編、Chapter 5“Design and Application of Prodrugs,”H.Bundgaard著、p.113-191(1991)、c)H.Bundgaard,Advanced Drug Delivery Reviews,8:1-38(1992)、d)H.Bundgaard,et al.,Journal of Pharmaceutical Sciences,77:285(1988)、ならびにe)N.Kakeya,et al.,Chem.Pharm. Bull.,32:692(1984)(これらの各々は、参照により本明細書に具体的に組み込まれる)を参照されたい。 For additional examples of prodrug derivatives, see, for example, a) Design of Prodrugs, edited by H. Bundgaard (Elsevier, 1985), and Methods in Enzymology, Vol. 42, pp. 309-396, edited by K. Widder, et al. (Academic Press, 1985); b) A Textbook of Drug Design and Development, edited by Krogsgaard-Larsen and H. Bundgaard, Chapter 5 "Design and Application of Prodrugs," by H. Bundgaard, p. 113-191 (1991); c) H. Bundgaard, Advanced Drug Delivery Reviews, 8:1-38 (1992); d) H. Bundgaard, et al., Journal of Pharmaceutical Sciences, 77:285 (1988); and e) N. Kakeya, et al., Chem. Pharm. Bull., 32:692 (1984), each of which is specifically incorporated herein by reference.
さらに、本発明は、本発明の化合物の代謝産物を提供する。本明細書で使用される場合、「代謝産物」とは、特定の化合物またはその塩の体内での代謝を通じて生成される生成物を指す。このような生成物は、例えば、投与された化合物の酸化、還元、加水分解、アミド化、脱アミド化、エステル化、脱エステル化、酵素切断などから生じ得る。 Additionally, the present invention provides metabolites of the compounds of the present invention. As used herein, "metabolite" refers to a product produced through metabolism in the body of a particular compound or salt thereof. Such products may result, for example, from oxidation, reduction, hydrolysis, amidation, deamidation, esterification, deesterification, enzymatic cleavage, and the like, of the administered compound.
代謝産物は、典型的には、本発明の化合物の放射標識された(例えば、14Cまたは3H)同位体を調製し、ラット、マウス、モルモット、サルなどの動物、またはヒトに、検出可能な用量(例えば、約0.5mg/kg超)で非経口投与し、代謝が生じるのに十分な時間(典型的には、約30秒~30時間)をかけ、その変換産物を尿、血液、または他の生体試料から単離することによって特定される。これらの生成物は、標識されているため、容易に単離される(その他は、代謝産物中で生存するエピトープを結合することができる抗体の使用によって単離される)。代謝産物構造は、例えば、MS、LC/MS、またはNMR分析によって、従来の様式で決定される。概して、代謝産物の分析は、当業者に周知である従来の薬物代謝研究と同様に行われる。代謝産物は、別途インビボで見出されない限り、本発明の化合物の治療投薬のための診断アッセイにおいて有用である。 Metabolites are typically identified by preparing radiolabeled (e.g., 14C or 3H) isotopes of the compounds of the invention, parenterally administering them to animals such as rats, mice, guinea pigs, monkeys, or humans at detectable doses (e.g., greater than about 0.5 mg/kg), allowing sufficient time for metabolism to occur (typically about 30 seconds to 30 hours), and isolating the conversion products from urine, blood, or other biological samples. These products are easily isolated because they are labeled (others are isolated by the use of antibodies capable of binding epitopes surviving in the metabolites). Metabolite structures are determined in conventional fashion, for example, by MS, LC/MS, or NMR analysis. In general, analysis of metabolites is performed similarly to conventional drug metabolism studies well known to those of skill in the art. Metabolites are useful in diagnostic assays for therapeutic dosing of the compounds of the invention, unless otherwise found in vivo.
「患者」または「対象」という用語は本明細書全体を通して、本開示による組成物を用いた予防的治療を含む治療が提供される動物、好ましくはヒトまたは家畜を記載するために使用される。例えばヒト患者などの特定の動物に特異的な感染症、状態または病態の治療に関し、患者という用語は、例えばイヌまたは猫などの家庭内動物、または例えばウマ、ウシ、ヒツジなどの農業用動物を含む特定の動物を指す。概して本開示において患者という用語は、別段の示唆が無い限り、または当該用語を使用している状況から暗示されない限り、ヒト患者を指す。 The terms "patient" or "subject" are used throughout this specification to describe an animal, preferably a human or a domestic animal, to which treatment, including prophylactic treatment, with a composition according to the present disclosure is provided. With respect to treatment of an infection, condition or pathology specific to a particular animal, e.g., a human patient, the term patient refers to the particular animal, including domestic animals, e.g., dogs or cats, or agricultural animals, e.g., horses, cows, sheep, etc. In general, in this disclosure, the term patient refers to a human patient, unless otherwise indicated or implied by the context in which the term is used.
「有効」という語は、その意図される用途の文脈の範囲内で使用される場合、意図される結果を生じさせる化合物、組成物、または構成要素の量を記載するために使用される。有効という用語は、その他すべての有効量または有効濃度の用語を含み、それらは本出願で別途記載または使用される。 The term "effective," when used within the context of its intended use, is used to describe an amount of a compound, composition, or component that produces an intended result. The term effective includes all other effective amount or effective concentration terms that are otherwise described or used in this application.
化合物および組成物
一態様では、本明細書は、フォンヒッペル・リンダウE3ユビキチンリガーゼ(VHL)結合部分(VLM)であるE3ユビキチンリガーゼ結合部分(「ULM」)を含む化合物を提供する。例示的実施形態において、ULMは、以下の構造に従い化学リンカー(L)を介して標的タンパク質結合部分(PTM)に連結される:
(A)PTM-L-ULM
式中、Lは結合または化学リンカー基であり、ULMはE3ユビキチンリガーゼ結合部分であり、そしてPTMは標的タンパク質結合部分である。本明細書に記載される化合物中の部分の数、および/または相対的な位置は、例示の目的でのみ提供される。当業者には理解されるように、本明細書に記載の化合物は、任意の望ましい数の各官能基部分を用いて、および/または各官能基部分の相対位置で合成することができる。
Compounds and Compositions In one aspect, the present disclosure provides compounds that include an E3 ubiquitin ligase binding moiety ("ULM") that is a von Hippel-Lindau E3 ubiquitin ligase (VHL) binding moiety (VLM). In an exemplary embodiment, the ULM is linked to a target protein binding moiety (PTM) via a chemical linker (L) according to the following structure:
(A) PTM-L-ULM
where L is a bond or chemical linker group, ULM is an E3 ubiquitin ligase binding moiety, and PTM is a target protein binding moiety. The number and/or relative positions of moieties in the compounds described herein are provided for illustrative purposes only. As will be appreciated by those skilled in the art, the compounds described herein can be synthesized with any desired number of each functional moiety and/or relative positions of each functional moiety.
別の態様において本開示は、標的タンパク質の分解を誘導することによるタンパク質活性の制御に有用な二官能性化合物または多官能性化合物(例えばPROTAC)を提供する。ある特定の実施形態では、化合物は、標的タンパク質(すなわち、タンパク質標的化部分または「PTM」)に結合する部分に、例えば、共有結合的に、直接的に、または間接的に結合されたVLMを含む。ある特定の実施形態では、VLMおよびPTMは、化学リンカー(L)を介して結合または結合される。VLMはVHLに結合し、PTMは標的タンパク質を認識し、それぞれの部分とそれらの標的との相互作用は、標的タンパク質をユビキチンリガーゼタンパク質に近接して配置することによって、標的タンパク質の分解を促進する。例示的な二官能性化合物は、以下のように示され得る:PTM-VLMとして表すことができる。 In another aspect, the disclosure provides bifunctional or multifunctional compounds (e.g., PROTACs) useful for controlling protein activity by inducing degradation of a target protein. In certain embodiments, the compounds include a VLM attached, e.g., covalently, directly, or indirectly, to a moiety that binds to the target protein (i.e., a protein targeting moiety or "PTM"). In certain embodiments, the VLM and PTM are attached or linked via a chemical linker (L). The VLM binds to the VHL, the PTM recognizes the target protein, and the interaction of each moiety with their target promotes degradation of the target protein by placing the target protein in close proximity to a ubiquitin ligase protein. An exemplary bifunctional compound can be depicted as follows: PTM-VLM.
特定の実施形態では、二官能性化合物は化学リンカー(「L」)をさらに含む。例えば、二官能性化合物は、以下のように示すことができる:PTM-L-VLM
式中、PTMは、タンパク質/ポリペプチド標的化部分であり、Lは、化学リンカーであり、VLMは、VHL結合部分である。
In certain embodiments, the bifunctional compound further comprises a chemical linker ("L"). For example, the bifunctional compound can be depicted as follows: PTM-L-VLM
where PTM is a protein/polypeptide targeting moiety, L is a chemical linker, and VLM is a VHL binding moiety.
ある特定の実施形態では、ULM(例えば、VLM)は、約200μM未満のIC50でE3ユビキチンリガーゼ(例えば、VHL)に活性を示すか、またはそれに結合する。IC50は、例えば、蛍光偏光アッセイなどの当分野で公知の任意の方法に従って決定することができる。 In certain embodiments, the ULM (e.g., VLM) exhibits activity against or binds to an E3 ubiquitin ligase (e.g., VHL) with an IC50 of less than about 200 μM. The IC50 can be determined according to any method known in the art, such as, for example, a fluorescence polarization assay.
ある特定の追加の実施形態では、本明細書に記載される二官能性化合物は、約100、50、10、1、0.5、0.1、0.05、0.01、0.005、0.001mM未満、または約100、50、10、1、0.5、0.1、0.05、0.01、0.005、0.001μM未満、または約100、50、10、1、0.5、0.1、0.05、0.01、0.005、0.001nM未満、または約100、50、10、1、0.5、0.1、0.05、0.01、0.005、0.001pM未満のIC50を有する活性を示す。 In certain additional embodiments, the bifunctional compounds described herein exhibit activity with an IC50 of less than about 100, 50, 10, 1, 0.5, 0.1, 0.05, 0.01, 0.005, 0.001 mM, or less than about 100, 50, 10, 1, 0.5, 0.1, 0.05, 0.01, 0.005, 0.001 μM, or less than about 100, 50, 10, 1, 0.5, 0.1, 0.05, 0.01, 0.005, 0.001 nM, or less than about 100, 50, 10, 1, 0.5, 0.1, 0.05, 0.01, 0.005, 0.001 pM.
特定の実施形態では、化合物が複数のULMを含む場合、それらULMは同一である。追加的な実施形態では、化合物は、複数のULM(例えばULM、ULM’)、直接、または化学リンカー(L)を介して、またはその両方でULMに結合される少なくとも1つのPTMを含む。特定の追加的な実施形態では、複数のULMを含む化合物は、複数のPTMをさらに含む。さらに追加的な実施形態では、PTMは同一であるか、または任意で異なっている。またさらなる実施形態では、PTMが異なる場合、各PTMは同じタンパク質標的に結合してもよく、または異なるタンパク質標的に特異的に結合してもよい。 In certain embodiments, when a compound comprises multiple ULMs, the ULMs are the same. In additional embodiments, a compound comprises multiple ULMs (e.g., ULM, ULM'), at least one PTM attached to the ULM directly, or via a chemical linker (L), or both. In certain additional embodiments, a compound comprising multiple ULMs further comprises multiple PTMs. In still additional embodiments, the PTMs are the same or optionally different. In still further embodiments, when the PTMs are different, each PTM may bind to the same protein target or may specifically bind to different protein targets.
特定の実施形態では、化合物は複数のULMおよび/または複数のULM’を含んでもよい。さらなる実施形態では、少なくとも2つの異なるULM、複数のULM、および/または複数のULM’を含む化合物は、直接、もしくは化学リンカーを介して、もしくはその両方を介してULMまたはULM’に結合された少なくとも1つのPTMをさらに含む。本明細書に記載の実施形態のいずれかにおいて、少なくとも2つの異なるULMを含む化合物は、複数のPTMをさらに含んでもよい。さらに追加的な実施形態では、PTMは同一であるか、または任意で異なっている。またさらなる実施形態では、PTMが異なる場合、各PTMは同じタンパク質標的に結合してもよく、または異なるタンパク質標的に特異的に結合してもよい。 In certain embodiments, the compound may include multiple ULMs and/or multiple ULMs'. In further embodiments, the compound including at least two different ULMs, multiple ULMs, and/or multiple ULMs' further includes at least one PTM attached to the ULM or ULM' either directly or via a chemical linker, or both. In any of the embodiments described herein, the compound including at least two different ULMs may further include multiple PTMs. In still further embodiments, the PTMs are the same or optionally different. In still further embodiments, when the PTMs are different, each PTM may bind to the same protein target or may specifically bind to different protein targets.
追加的実施形態では、本明細書は、その鏡像異性体、ジアステレオマー、溶媒和物および多形体を含み、その薬学的に許容可能な塩型、例えば酸塩型および塩基塩型を含む、本明細書に記載の化合物を提供する。 In additional embodiments, the present disclosure provides compounds described herein, including their enantiomers, diastereomers, solvates, and polymorphs, including their pharma- ceutically acceptable salt forms, e.g., acid salt forms and base salt forms.
例示的なVLM
ある特定の実施形態では、本明細書に記載される化合物は、E3ユビキチンリガーゼ、例えば、フォンヒッペル・リンダウE3ユビキチンリガーゼに結合するための手段を含む。ある特定の実施形態では、ULMは、VLMであり、かつ以下のULM-a群から選択される化学構造を含み、
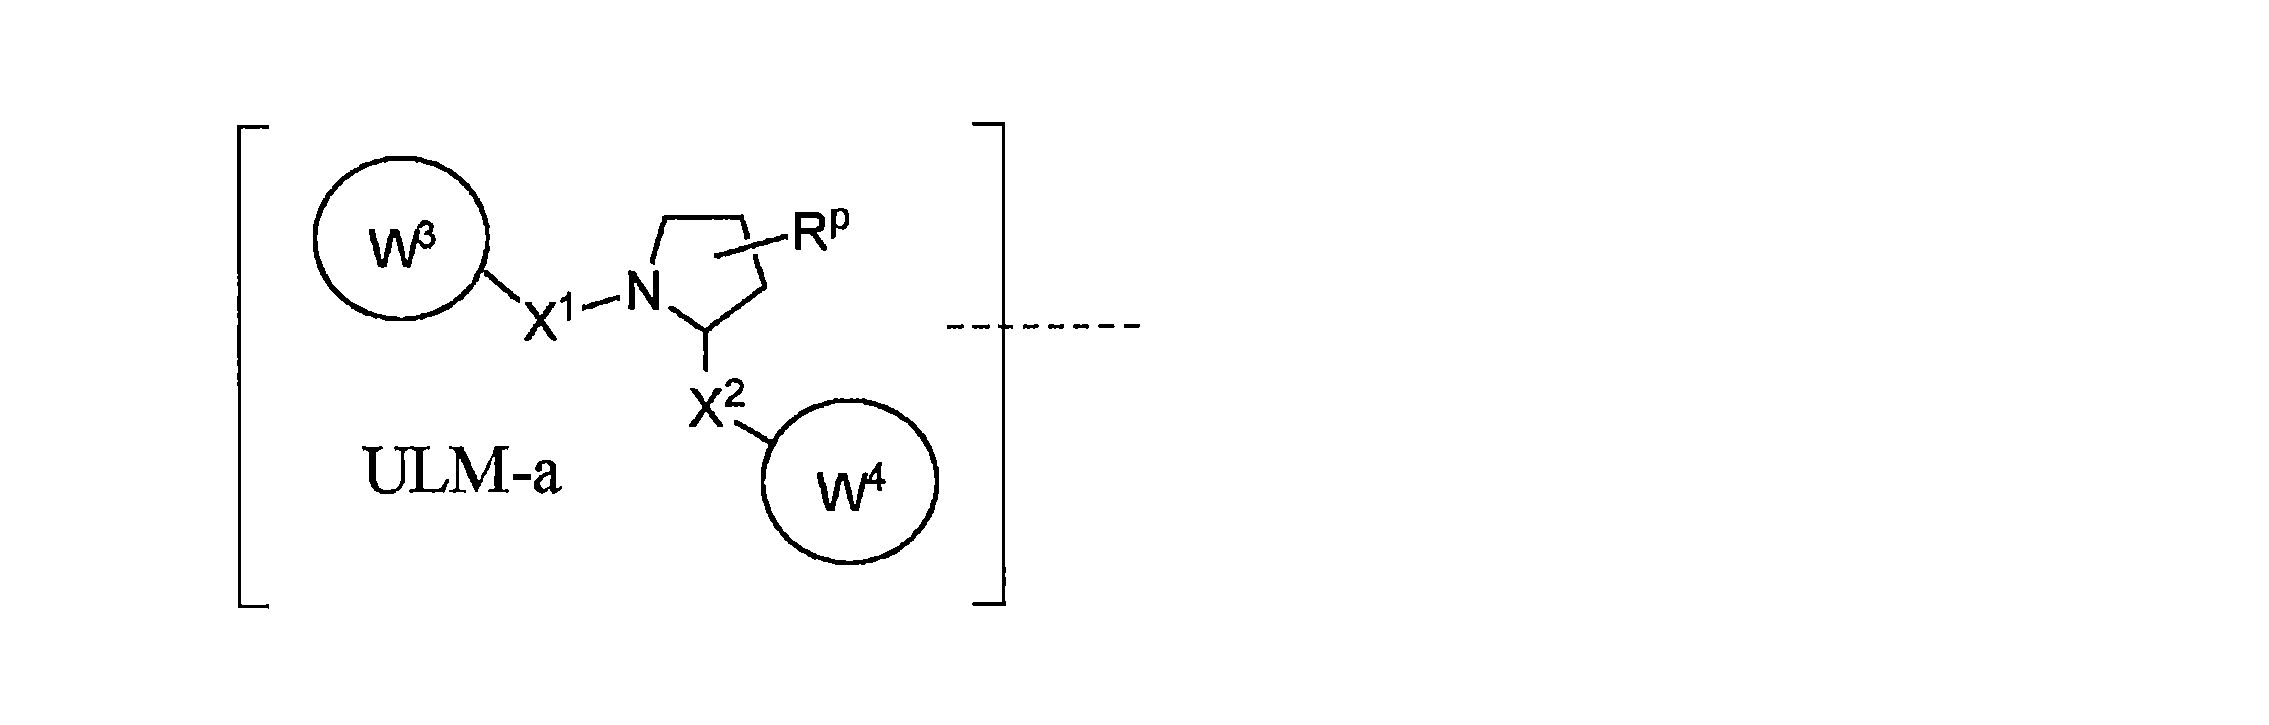
破線は、少なくとも1つのPTM、別のULMもしくはVLM(VLM’)、または少なくとも1つのPTM、ULM’、もしくはVLM’をリンカーの他方の端部に結合させる化学リンカー部分の結合を示し、
式ULM-aのX1、X2はそれぞれ独立して、結合、O、NRY3、CRY3RY4、C=O、C=S、SO、およびSO2の群から選択され、
式ULM-aのRY3、RY4は、各々独立して、H、1つ以上のハロで任意に置換された直鎖または分岐鎖C1-6アルキル、任意に置換された(例えば、0~3個のRP基で任意に置換された)C1-6アルコキシルからなる群から選択され、
式ULM-aのRPは、0、1、2、または3個の基であり、各々が独立して、H、ハロ、-OH、C1-3アルキル、C=O、アルキル、アルコキシ、またはそれらの組み合わせの群から選択され、
式ULM-aのW3は、任意に置換されたT、任意に置換された-T-N(R1aR1b)X3、任意に置換された-T-N(R1aR1b)、任意に置換された-T-アリール、任意に置換された-T-ヘテロアリール、任意に置換されたT-ビヘテロアリール、任意に置換された-T-ヘテロシクリル、任意に置換された-T-ビヘテロシクリル、任意に置換された-NR1-T-アリール、任意に置換された-NR1-T-ヘテロアリール、または任意に置換された-NR1-T-ヘテロシクリルの群から選択され、
式ULM-aのX3は、C=O、R1、R1a、R1bであり、
R1、R1a、R1bの各々は、独立して、H、1個以上のハロで任意に置換された直鎖もしくは分岐鎖C1-C6アルキル基、または-OH基、RY3C=O、RY3C=S、RY3SO、RY3SO2、N(RY3RY4)C=O、N(RY3RY4)C=S、N(RY3RY4)SO、およびN(RY3RY4)SO2からなる群から選択され、
式ULM-aのTは、任意に置換されたアルキル、-(CH2)n-基、任意に置換された直鎖もしくは分岐鎖-(CH2)n-O-C1-C6アルキル、または任意に置換された-(CH2)n-O-ヘテロシクリルの群から選択され、式中、メチレン基の各々は、ハロゲン、メチル、1個以上のハロゲンで任意に置換された直鎖もしくは分岐鎖C1-C6アルキル基、または-OH基、任意に置換されたアミノ酸側鎖、または任意に置換されたヘテロシクリルの群から選択される1個または2個の置換基で置換され、
式ULM-aのW4は、任意に置換された-NR1-T-アリールであり、式中、アリール基は、任意に置換された5~6員ヘテロアリール、または任意に置換されたアリール、任意に置換された-NR1-T-ヘテロアリール基であり得、式中、ヘテロアリールは、任意に置換されたアリール、または任意に置換されたヘテロアリール、または任意に置換された-NR1-T-ヘテロシクリルで任意に置換され、式中、-NR1は、X2に共有結合し、R1がHまたはCH3、好ましくはHである。
Exemplary VLMs
In certain embodiments, the compounds described herein comprise a means for binding to an E3 ubiquitin ligase, such as a von Hippel-Lindau E3 ubiquitin ligase. In certain embodiments, the ULM is a VLM and comprises a chemical structure selected from the ULM-a group:
The dashed lines indicate the attachment of at least one PTM, another ULM or VLM (VLM'), or a chemical linker moiety that attaches at least one PTM, ULM', or VLM' to the other end of the linker;
Each of X1 and X2 in formula ULM-a is independently selected from the group consisting of a bond, O, NR, CRY, C=O, C=S, SO, and SO;
Each of RY3 and RY4 in formula ULM-a is independently selected from the group consisting of H, straight or branched C1-6 alkyl optionally substituted with one or more halo, and optionally substituted (e.g., optionally substituted with 0-3 RP groups) C1-6 alkoxyl;
R of formula ULM-a is 0, 1, 2, or 3 groups, each independently selected from the group of H, halo, —OH, C1-3 alkyl, C═O, alkyl, alkoxy, or combinations thereof;
W3 of formula ULM-a is selected from the group of optionally substituted T, optionally substituted -T-N(R1aR1b)X3, optionally substituted -T-N(R1aR1b), optionally substituted -T-aryl, optionally substituted -T-heteroaryl, optionally substituted T-biheteroaryl, optionally substituted -T-heterocyclyl, optionally substituted -T-biheterocyclyl, optionally substituted -NR1-T-aryl, optionally substituted -NR1-T-heteroaryl, or optionally substituted -NR1-T-heterocyclyl;
X3 in formula ULM-a is C=O, R1, R1a, or R1b;
Each of R1, R1a, and R1b is independently selected from the group consisting of H, a straight or branched C1-C6 alkyl group optionally substituted with one or more halo, or an -OH group, RY3C=O, RY3C=S, RY3SO, RY3SO2, N(RY3RY4)C=O, N(RY3RY4)C=S, N(RY3RY4)SO, and N(RY3RY4)SO2;
T of formula ULM-a is selected from the group of optionally substituted alkyl, -(CH2)n- group, optionally substituted linear or branched -(CH2)n-O-C1-C6 alkyl, or optionally substituted -(CH2)n-O-heterocyclyl, wherein each of the methylene groups is substituted with 1 or 2 substituents selected from the group of halogen, methyl, linear or branched C1-C6 alkyl group optionally substituted with one or more halogens, or -OH group, optionally substituted amino acid side chain, or optionally substituted heterocyclyl;
W4 in formula ULM-a is an optionally substituted -NR1-T-aryl, where the aryl group can be an optionally substituted 5-6 membered heteroaryl, or an optionally substituted aryl, an optionally substituted -NR1-T-heteroaryl group, where the heteroaryl is optionally substituted with an optionally substituted aryl, or an optionally substituted heteroaryl, or an optionally substituted -NR1-T-heterocyclyl, where -NR1 is covalently bonded to X2 and R1 is H or CH3, preferably H.
ある特定の実施形態では、RPは、エステルまたはエーテル結合を含む、プロドラッグを形成するように修飾される。 In certain embodiments, RP is modified to form a prodrug, including an ester or ether bond.
本明細書に記載される任意の態様または実施形態では、Tは、任意に置換されたアルキル、-(CH2)n-基の群から選択され、式中、メチレン基の各々は、ハロゲン、メチル、任意に置換されたアルコキシ、1個以上のハロゲンで任意に置換された直鎖もしくは分岐鎖C1-C6アルキル基、C(O)NR1R1a、またはNR1R1aの群から選択される1個または2個の置換基で任意に置換されるか、あるいはR1およびR1aが、結合されて、任意に置換されたヘテロシクリル、または-OH基、または任意に置換されたアミノ酸側鎖を形成し、nが、0~6、多くの場合0、1、2、または3、好ましくは0または1である。 In any aspect or embodiment described herein, T is selected from the group of an optionally substituted alkyl, -(CH2)n- group, where each methylene group is optionally substituted with one or two substituents selected from the group of halogen, methyl, optionally substituted alkoxy, linear or branched C1-C6 alkyl groups optionally substituted with one or more halogens, C(O)NR1R1a, or NR1R1a, or R1 and R1a are joined to form an optionally substituted heterocyclyl, or -OH group, or an optionally substituted amino acid side chain, and n is 0-6, often 0, 1, 2, or 3, preferably 0 or 1.
本明細書に記載される任意の態様または実施形態では、式ULM-aのW4は、
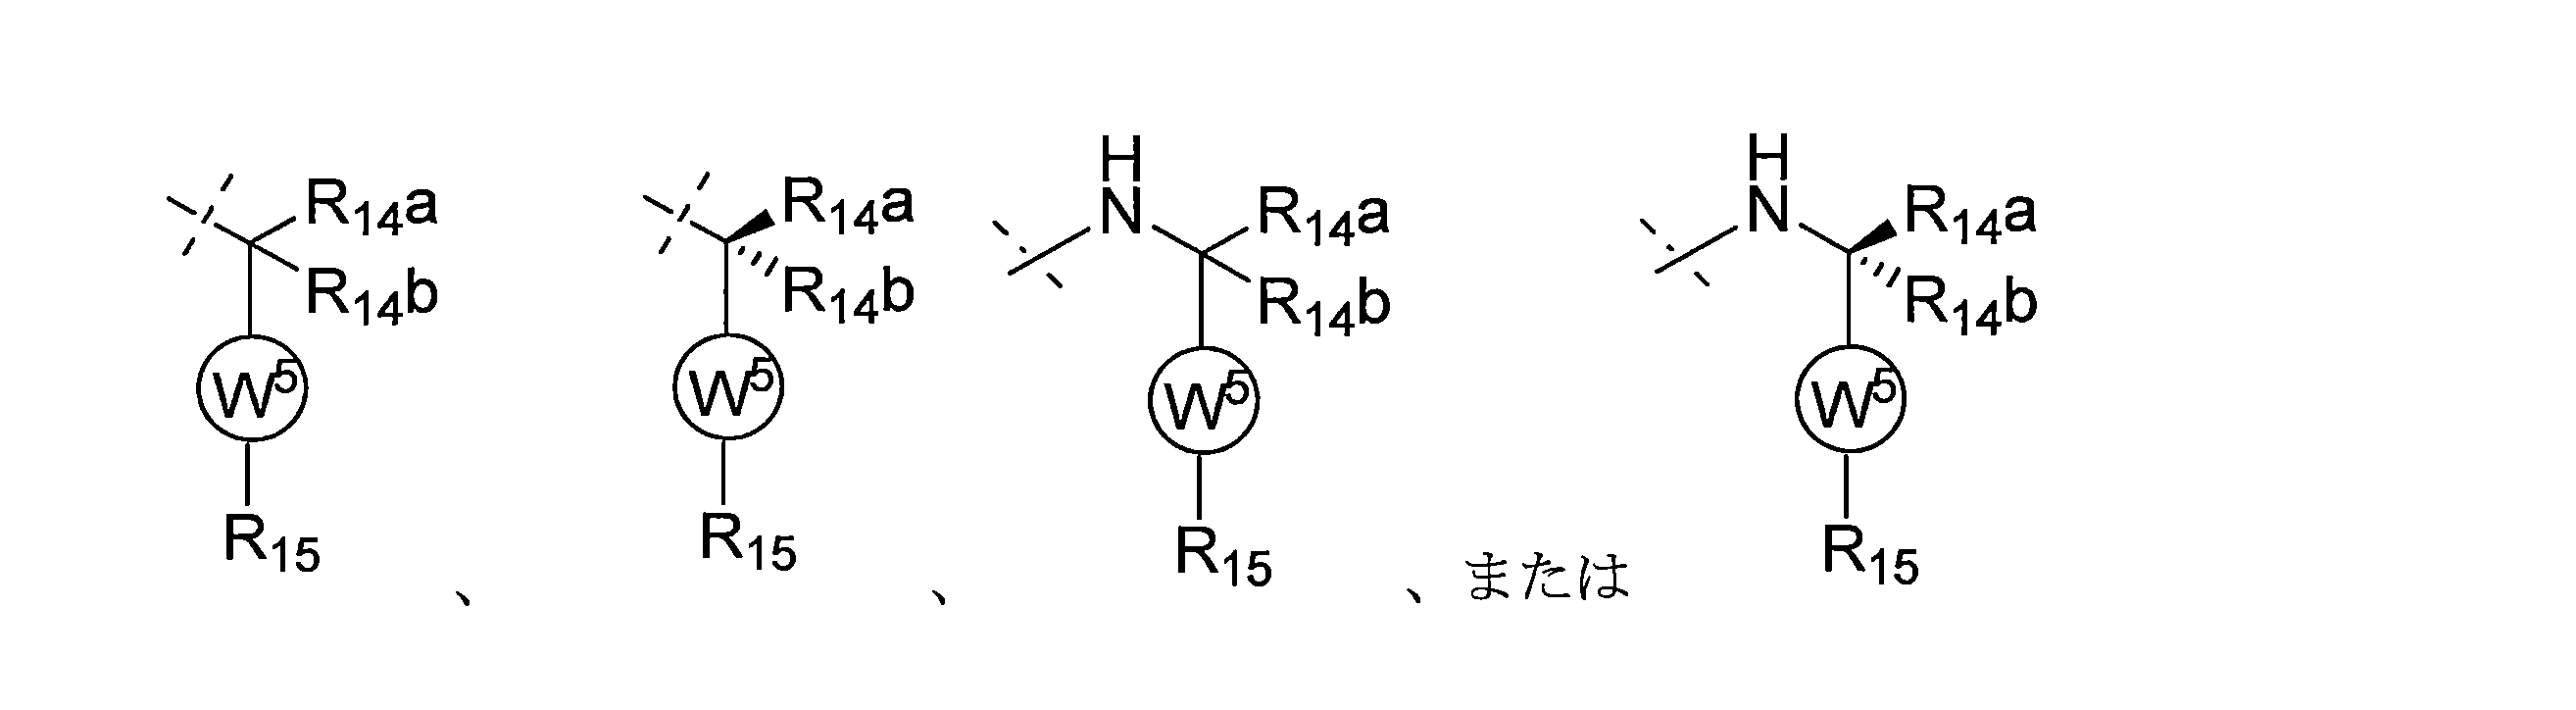
実施形態のいずれかでは、式ULM-aのW5は、任意に置換されたフェニル、任意に置換されたナプチル、または任意に置換された5~10員ヘテロアリールの群から選択され、
式ULM-aのR15は、H、ハロゲン、CN、C≡CH、OH、NO2、NR14aR14b、OR14a、CONR14aR14b、NR14aCOR14b、SO2NR14aR14b、NR14aSO2R14b、任意に置換されたアルキル、任意に置換されたハロアルキル、任意に置換されたハロアルコキシ、任意に置換されたアリール、任意に置換されたヘテロアリール、任意に置換されたシクロアルキル、または任意に置換されたヘテロシクリルの群から選択される。
In any of the embodiments, W5 of formula ULM-a is selected from the group of optionally substituted phenyl, optionally substituted naphthyl, or optionally substituted 5-10 membered heteroaryl;
R15 of formula ULM-a is selected from the group of H, halogen, CN, C≡CH, OH, NO2, NR14aR14b, OR14a, CONR14aR14b, NRR14b, SO2NR14aR14b, NR14aSO2R14b, optionally substituted alkyl, optionally substituted haloalkyl, optionally substituted haloalkoxy, optionally substituted aryl, optionally substituted heteroaryl, optionally substituted cycloalkyl, or optionally substituted heterocyclyl.
追加の実施形態では、本開示における使用のためのW4置換基はまた、本明細書に開示される特定の化合物中に見出されるW4置換基を特異的に含む(かつ、開示される特定の化合物に限定されない)。これらW4置換基の各々は、任意の数のW3置換基と併せて使用されてもよく、それらも本明細書に開示されている。 In additional embodiments, W4 substituents for use in the present disclosure also specifically include (and are not limited to) the W4 substituents found in the particular compounds disclosed herein. Each of these W4 substituents may be used in conjunction with any number of W3 substituents, which are also disclosed herein.
ある特定の追加の実施形態では、ULM-aは、ピロリジン部分中の0~3個のRP基によって任意に置換される。各RPは独立して、H、ハロ、-OH、C1-3アルキル、C=Oである。 In certain additional embodiments, ULM-a is optionally substituted with 0-3 RP groups in the pyrrolidine moiety. Each RP is independently H, halo, -OH, C1-3 alkyl, C=O.
本明細書に記載される実施形態のいずれかでは、式ULM-aのW3、W4は、独立して、1つ以上のPTM基に結合しているリンカーに共有結合することができ、
および式中、破線は、少なくとも1つのPTM、別のULM(ULM’)、または少なくとも1つのPTMもしくはULM’もしくはその両方をULMに結合する化学リンカー部分の結合部位を示す。
In any of the embodiments described herein, W3, W4 of formula ULM-a can be independently covalently attached to a linker that is attached to one or more PTM groups;
and wherein the dashed lines indicate the site of attachment of at least one PTM, another ULM (ULM'), or a chemical linker moiety linking at least one PTM or ULM' or both to a ULM.
特定の実施形態では、ULMはVHLであり、以下の構造によって表される:
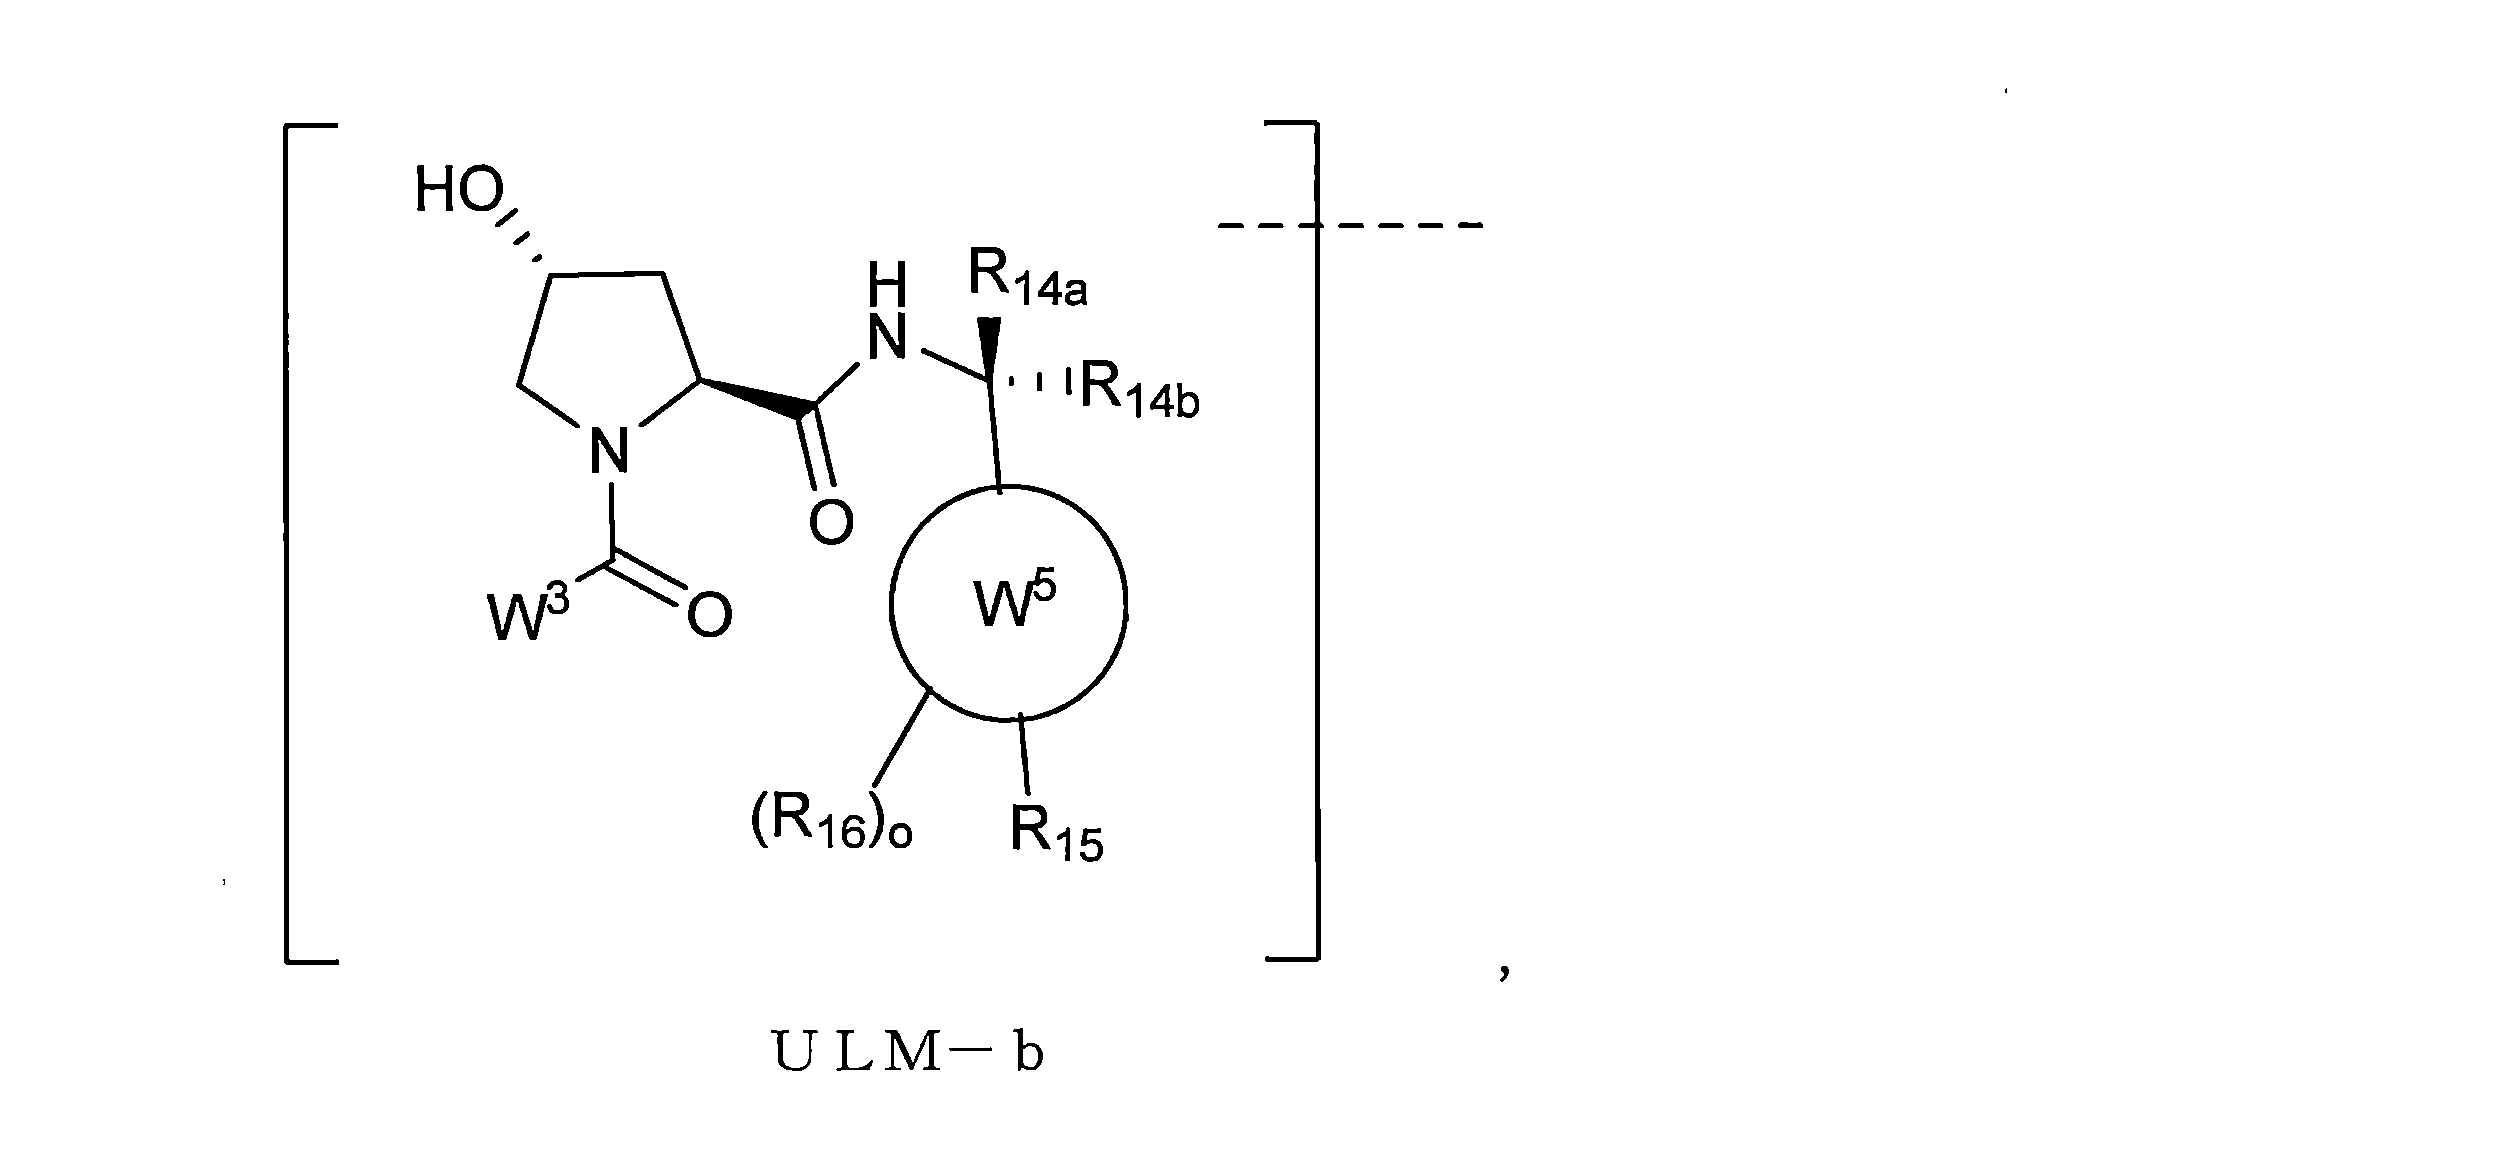
式ULM-bのW3は、任意に置換されたアリール、任意に置換されたヘテロアリール、または

式ULM-bのR9およびR10は、独立して、水素、任意に置換されたアルキル、任意に置換されたシクロアルキル、任意に置換されたヒドロキシアルキル、任意に置換されたヘテロアリール、もしくはハロアルキルであるか、またはR9、R10、およびそれらが結合する炭素原子が、任意に置換されたシクロアルキルを形成し、
式ULM-bのR11は、任意に置換されたヘテロシクリル、任意に置換されたアルコキシ、任意に置換されたヘテロアリール、任意に置換されたアリール、
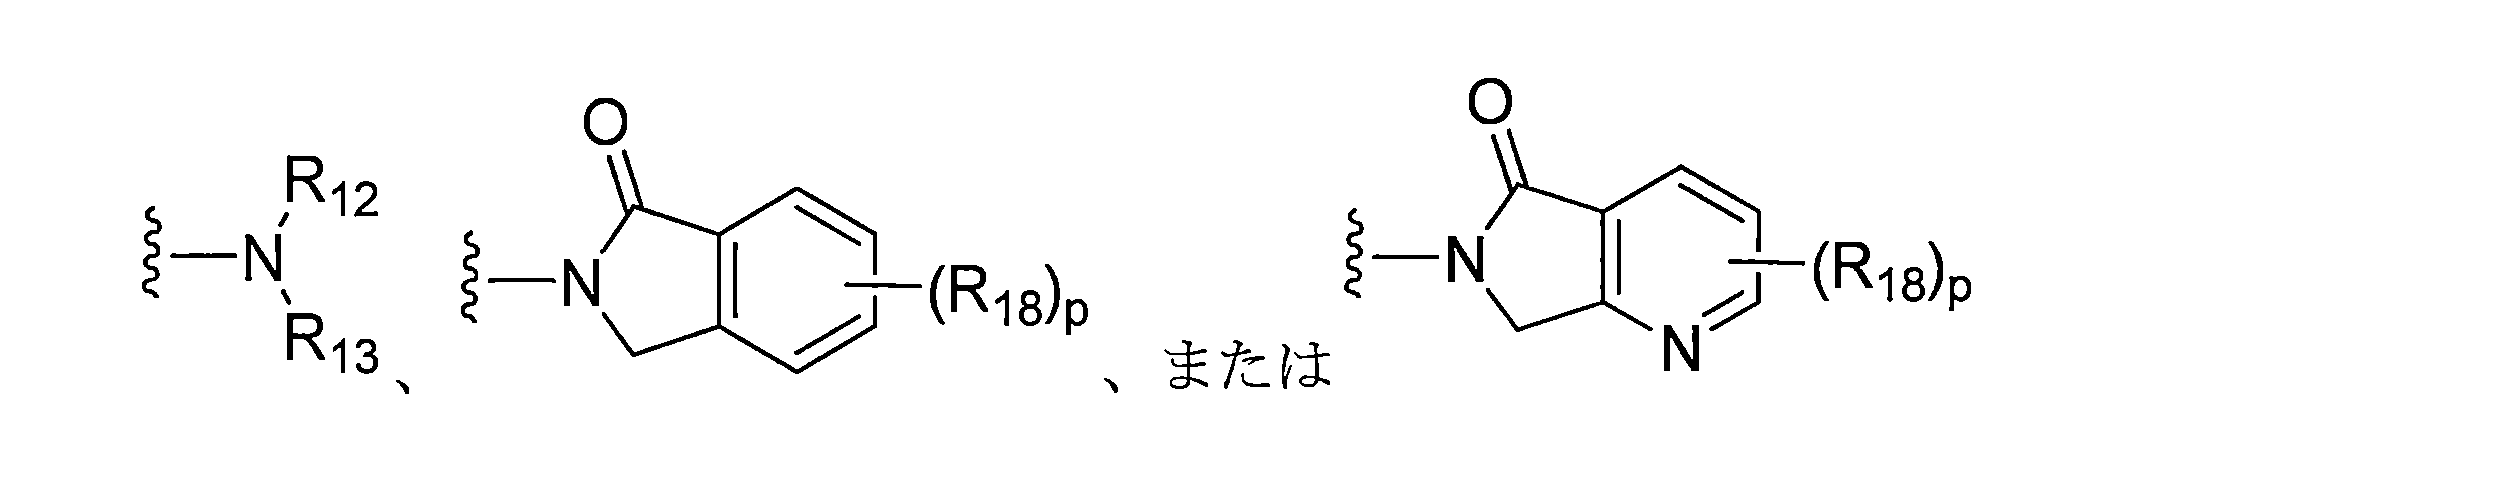
式ULM-bのR12は、Hまたは任意に置換されたアルキルの群から選択され、
式ULM-bのR13は、H、任意に置換されたアルキル、任意に置換されたアルキルカルボニル、任意に置換された(シクロアルキル)アルキルカルボニル、任意に置換されたアラルキルカルボニル、任意に置換されたアリールカルボニル、任意に置換された(ヘテロシクリル)カルボニル、または任意に置換されたアラルキルの群から選択され、
式ULM-bのR14a、R14bは、各々独立して、H、ハロアルキル(例えば、フルオロアルキル)、任意に置換されたアルキル、任意に置換されたアルコキシ、アミノメチル、アルキルアミノメチル、アルコキシメチル、任意に置換されたヒドロキシルアルキル、任意に置換されたアルキルアミン、任意に置換されたアミド、任意に置換されたアルキル-アミド、任意に置換されたアルキル-シアノ、任意に置換されたアルキル-ホスフェート、任意に置換されたヘテロアルキル、任意に置換されたアルキル-ヘテロシクロアルキル、任意に置換されたアルコキシ-ヘテロシクロアルキル、COR26、アルキル-COR26、CONR27aR27b、CH2NHCOR26、もしくは(CH2)N(CH3)COR26の群から選択され、R14aおよびR14bのうちの他方は、Hであるか、またはR14a、R14bは、それらが結合している炭素原子と一緒になって、任意に置換された3~6員シクロアルキル、ヘテロシクロアルキ、スピロシクロアルキル、もしくはスピロヘテロシクリルを形成し、スピロヘテロシクリルは、エポキシドまたはアジリジンではない。
式ULM-bのW5は、フェニル、ナプチル、または5~10員ヘテロアリールの群から選択され、
式ULM-bのR15は、H、ハロゲン、CN、C≡CH、OH、NO2、NR27aR27b、OR27a、CONR27aR27b、NR27aCOR27b、SO2NR27aR27b、NR27aSO2R27b、任意に置換されたアルキル、任意に置換されたハロアルキル、任意に置換されたハロアルコキシ、任意に置換されたアリール、任意に置換されたヘテロアリール、任意に置換されたシクロアルキル、または任意に置換されたヘテロシクリルの群から選択され、
式ULM-bの各R16は、独立して、ハロ、CN、任意に置換されたアルキル、任意に置換されたアルキルアミン、任意に置換されたハロアルキル、ヒドロキシ、または任意に置換されたハロアルコキシの群から選択され、
式ULM-bのoは、0、1、2、3、または4であり、
式ULM-bのR18は、H、ハロ、任意に置換されたアルコキシ、シアノ、任意に置換されたアルキル、ハロアルキル、ハロアルコキシまたはリンカーの群から独立して選択され、
各R26は、独立して、H、OH、任意に置換されたアルキル、またはNR27aR27bから選択され、
各R27aおよびR27bは、独立して、H、任意に置換されたアルキル、任意に置換された3~5員シクロアルキルであるか、またはR27aおよびR27bが、それらが結合している窒素原子と一緒になって、4~6員ヘテロシクリルを形成し、
式ULM-bのpは、0、1、2、3、または4であり、および式中、破線は、少なくとも1つのPTM、別のULM(ULM’)、または少なくとも1つのPTMもしくはULM’もしくはその両方をULMに結合する化学リンカー部分の結合部位を示す。
In certain embodiments, the ULM is VHL and is represented by the following structure:
W3 of formula ULM-b is optionally substituted aryl, optionally substituted heteroaryl, or
R9 and R10 of formula ULM-b are independently hydrogen, optionally substituted alkyl, optionally substituted cycloalkyl, optionally substituted hydroxyalkyl, optionally substituted heteroaryl, or haloalkyl, or R9, R10 and the carbon atom to which they are attached form an optionally substituted cycloalkyl;
R11 of formula ULM-b is optionally substituted heterocyclyl, optionally substituted alkoxy, optionally substituted heteroaryl, optionally substituted aryl,
R12 of formula ULM-b is selected from the group of H or optionally substituted alkyl;
R13 of formula ULM-b is selected from the group: H, optionally substituted alkyl, optionally substituted alkylcarbonyl, optionally substituted (cycloalkyl)alkylcarbonyl, optionally substituted aralkylcarbonyl, optionally substituted arylcarbonyl, optionally substituted (heterocyclyl)carbonyl, or optionally substituted aralkyl;
R14a and R14b in formula ULM-b are each independently H, haloalkyl (e.g., fluoroalkyl), optionally substituted alkyl, optionally substituted alkoxy, aminomethyl, alkylaminomethyl, alkoxymethyl, optionally substituted hydroxylalkyl, optionally substituted alkylamine, optionally substituted amido, optionally substituted alkyl-amido, optionally substituted alkyl-cyano, optionally substituted alkyl-phosphate, optionally substituted heteroalkyl, optionally substituted alkyl-heterocycloalkyl, any wherein the other of R14a and R14b is H, or R14a, R14b together with the carbon atom to which they are attached form an optionally substituted 3- to 6-membered cycloalkyl, heterocycloalkyl, spirocycloalkyl, or spiroheterocyclyl, and wherein the spiroheterocyclyl is not an epoxide or an aziridine.
W5 of formula ULM-b is selected from the group of phenyl, naphthyl, or 5-10 membered heteroaryl;
R15 of formula ULM-b is selected from the group H, halogen, CN, C≡CH, OH, NO2, NR27aR27b, OR27a, CONR27aR27b, NR27aCOR27b, SO2NR27aR27b, NR27aSO2R27b, optionally substituted alkyl, optionally substituted haloalkyl, optionally substituted haloalkoxy, optionally substituted aryl, optionally substituted heteroaryl, optionally substituted cycloalkyl, or optionally substituted heterocyclyl;
Each R16 of formula ULM-b is independently selected from the group of halo, CN, optionally substituted alkyl, optionally substituted alkylamine, optionally substituted haloalkyl, hydroxy, or optionally substituted haloalkoxy;
In the formula ULM-b, o is 0, 1, 2, 3, or 4;
R18 of formula ULM-b is independently selected from the group of H, halo, optionally substituted alkoxy, cyano, optionally substituted alkyl, haloalkyl, haloalkoxy or a linker;
each R26 is independently selected from H, OH, optionally substituted alkyl, or NR27aR27b;
each R27a and R27b is independently H, optionally substituted alkyl, optionally substituted 3- to 5-membered cycloalkyl, or R27a and R27b together with the nitrogen atom to which they are attached form a 4- to 6-membered heterocyclyl;
In formula ULM-b, p is 0, 1, 2, 3, or 4, and wherein the dashed lines indicate the site of attachment of at least one PTM, another ULM (ULM'), or a chemical linker moiety linking at least one PTM or ULM' or both to a ULM.
ある特定の実施形態では、式ULM-bのR15は、

特定の実施形態では、式ULM-bのR17は、メチル、エチル、イソプロピルおよびシクロプロピルの群から選択される。 In certain embodiments, R17 of formula ULM-b is selected from the group consisting of methyl, ethyl, isopropyl, and cyclopropyl.
特定の追加的実施形態では、式ULM-bのR15は、以下からなる群から選択される:
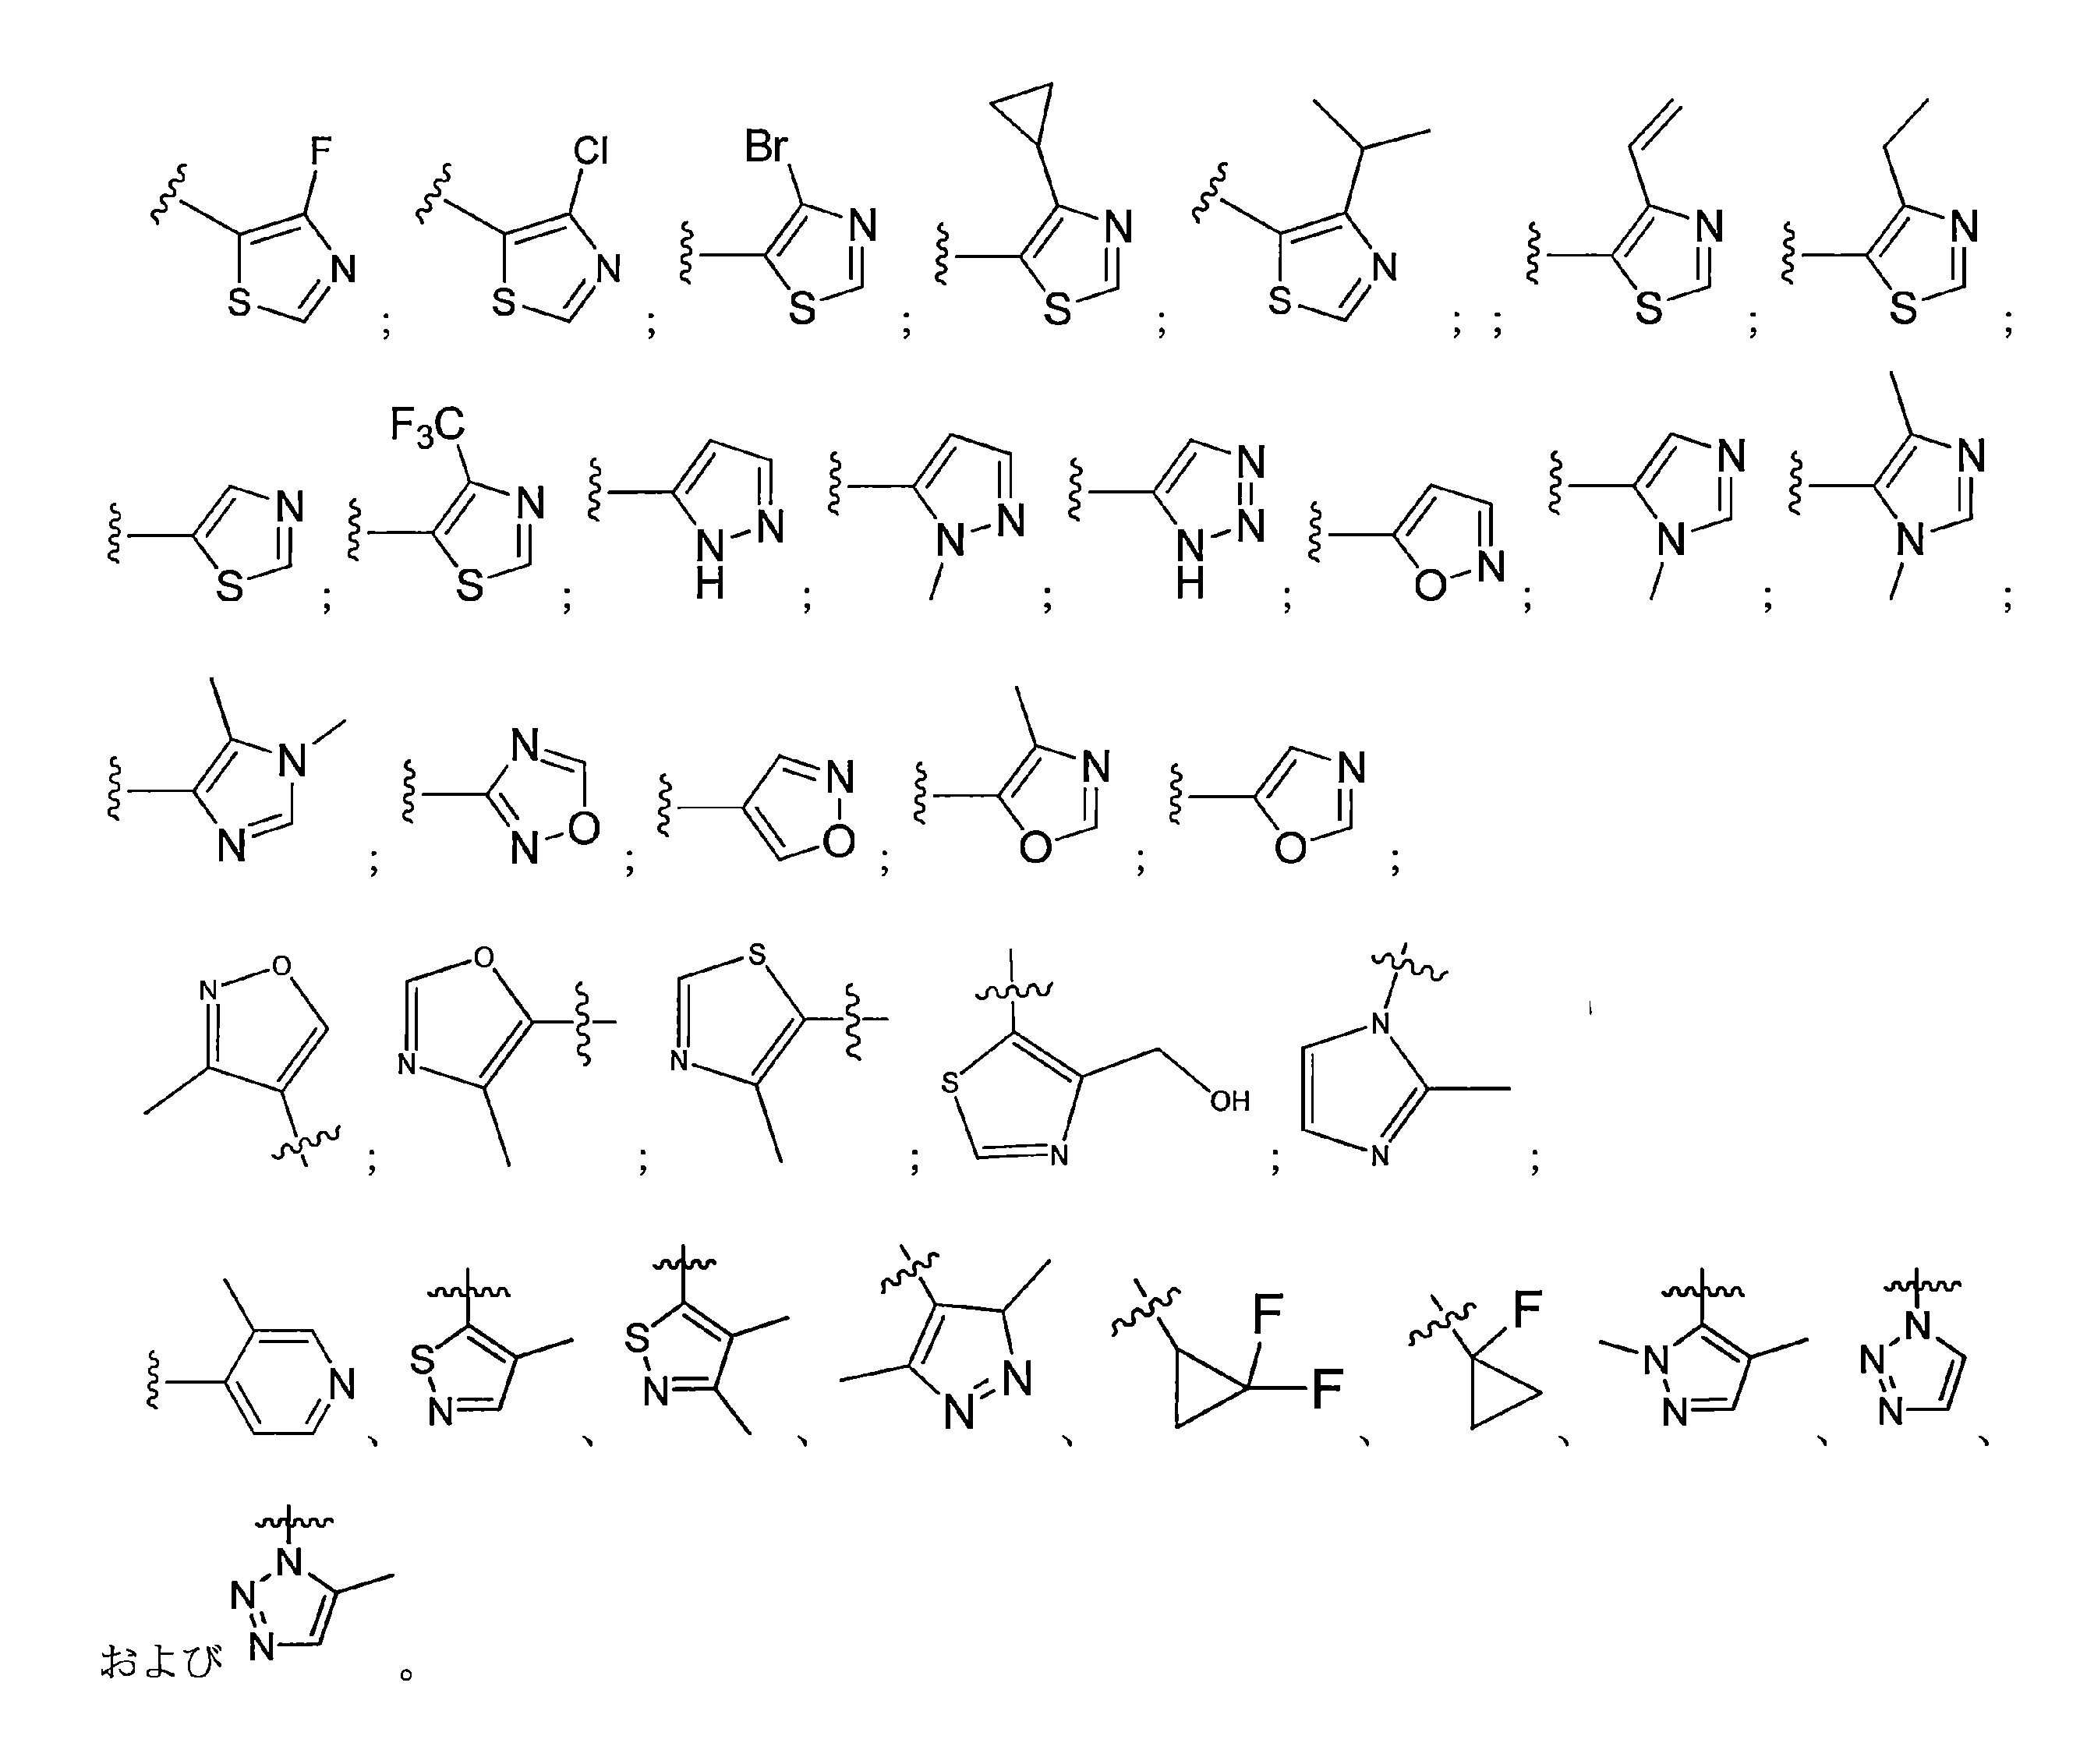
特定の実施形態では、式ULM-bのR11は、以下からなる群から選択される:

本明細書に記載される任意の態様または実施形態では、ULM-bのR14a、R14bは、各々独立して、H、任意に置換されたハロアルキル、任意に置換されたアルキル、任意に置換されたアルコキシ、任意に置換されたヒドロキシルアルキル、任意に置換されたアルキルアミン、任意に置換されたアミド、任意に置換されたアルキル-アミド、任意に置換されたアルキル-シアノ、任意に置換されたアルキル-ホスフェート、任意に置換されたヘテロルキル、任意に置換されたアルキル-ヘテロシクロアルキル、任意に置換されたアルコキシ-ヘテロシクロアルキル、COR26、アルキル-COR26、CH2OR30、CH2NHR30、CH2NCH3R30、CONR27aR27b、CH2CONR27aR27b、CH2NHCOR26、もしくはCH2NCH3COR26の群から選択され、R14aおよびR14bのうちの他方は、Hであるか、またはR14a、R14bは、それらが結合している炭素原子と一緒になって、任意に置換された3~6員シクロアルキル、ヘテロシクロアルキル、スピロシクロアルキル、もしくはスピロヘテロシクリルを形成し、スピロヘテロシクリルは、エポキシドまたはアジリジンではなく、該スピロシクロアルキルもしくはスピロヘテロシクリル自体は、アルキル、ハロアルキル、または-COR33で任意に置換され、式中、R33は、アルキルもしくはハロアルキルであり、
式中、R30は、H、アルキル、アルキニルアルキル、シクロアルキル、ヘテロシクロアルキル、シクロアルキルアルキル、ヘテロシクロアルキルアルキル、アリールアルキル、またはさらに任意に置換されたヘテロアリールアルキルから選択され、R26およびR27は、上記のとおりである。
In any aspect or embodiment described herein, R14a, R14b of ULM-b are each independently H, optionally substituted haloalkyl, optionally substituted alkyl, optionally substituted alkoxy, optionally substituted hydroxylalkyl, optionally substituted alkylamine, optionally substituted amide, optionally substituted alkyl-amide, optionally substituted alkyl-cyano, optionally substituted alkyl-phosphate, optionally substituted heteroalkyl, optionally substituted alkyl-heterocycloalkyl, optionally substituted alkoxy-heterocycloalkyl, COR26, alkyl-COR26, CH2OR30, CH2NHR30, CH2NCH3R3 0, CONR27aR27b, CH2CONR27aR27b, CH2NHCOR26, or CH2NCH3COR26; the other of R14a and R14b is H, or R14a, R14b together with the carbon atom to which they are attached form an optionally substituted 3-6 membered cycloalkyl, heterocycloalkyl, spirocycloalkyl, or spiroheterocyclyl, which spiroheterocyclyl is not an epoxide or aziridine, and said spirocycloalkyl or spiroheterocyclyl is itself optionally substituted with alkyl, haloalkyl, or -COR33, where R33 is alkyl or haloalkyl;
wherein R30 is selected from H, alkyl, alkynylalkyl, cycloalkyl, heterocycloalkyl, cycloalkylalkyl, heterocycloalkylalkyl, arylalkyl, or further optionally substituted heteroarylalkyl; and R26 and R27 are as defined above.
本明細書に記載される任意の態様または実施形態では、式ULM-bのR15は、H、ハロゲン、CN、C≡CH、OH、NO2、NR27aR27b、OR27a、CONR27aR27b、NR27aCOR27b、SO2NR27aR27b、NR27aSO2R27b、任意に置換されたアルキル、任意に置換されたハロアルキル(例えば、任意に置換されたフルオロアルキル)、任意に置換されたハロアルコキシ、任意に置換されたアリール、任意に置換されたヘテロアリール、任意に置換されたシクロアルキル、または任意に置換されたヘテロシクリルから選択され、式中、該アリール、ヘテロアリール、シクロアルキル、およびヘテロシクロアルキルの任意の置換は、CH2OR30、CH2NHR30、CH2NCH3R30、CONR27aR27b、CH2CONR27aR27b、CH2NHCOR26、CH2NCH3COR26、または

本明細書に記載される任意の態様または実施形態では、式ULM-bのR14a、R14bは、各々独立して、H、任意に置換されたハロアルキル、任意に置換されたアルキル、CH2OR30、CH2NHR30、CH2NCH3R30、CONR27aR27b、CH2CONR27aR27b、CH2NHCOR26、もしくはCH2NCH3COR26の群から選択され、R14aおよびR14bのうちの他方は、Hであるか、またはR14a、R14bは、それらが結合している炭素原子と一緒になって、任意に置換された3~6員スピロシクロアルキル、もしくはスピロヘテロシクリルを形成し、スピロヘテロシクリルは、エポキシドまたはアジリジンではなく、該スピロシクロアルキルもしくはスピロヘテロシクリル自体は、アルキル、ハロアルキル、または-COR33で任意に置換され、式中、R33は、アルキルもしくはハロアルキルであり、式中、R30は、H、アルキル、アルキニルアルキル、シクロアルキル、ヘテロシクロアルキル、シクロアルキルアルキル、ヘテロシクロアルキルアルキル、アリールアルキル、またはさらに任意に置換されたヘテロアリールアルキルから選択され、
式ULM-bのR15は、H、ハロゲン、CN、C≡CH、OH、NO2、NR27aR27b、OR27a、CONR27aR27b、NR27aCOR27b、SO2NR27aR27b、NR27aSO2R27b、任意に置換されたアルキル、任意に置換されたハロアルキル、任意に置換されたハロアルコキシ、任意に置換されたアリール、任意に置換されたヘテロアリール、任意に置換されたシクロアルキル、または任意に置換されたヘテロシクリルから選択され、式中、該アリール、ヘテロアリール、シクロアルキル、およびヘテロシクロアルキルの任意の置換は、CH2OR30、CH2NHR30、CH2NCH3R30、CONR27aR27b、CH2CONR27aR27b、CH2NHCOR26、CH2NCH3COR26、または
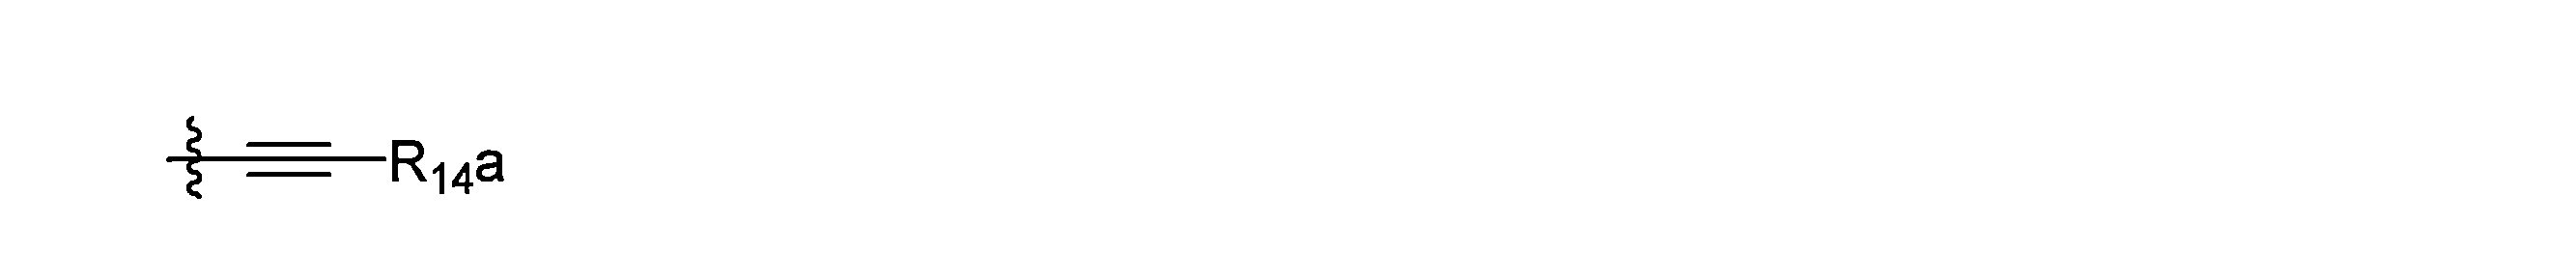
R15 of formula ULM-b is selected from H, halogen, CN, C≡CH, OH, NO2, NR27aR27b, OR27a, CONR27aR27b, NR27aCOR27b, SO2NR27aR27b, NR27aS02R27b, optionally substituted alkyl, optionally substituted haloalkyl, optionally substituted haloalkoxy, optionally substituted aryl, optionally substituted heteroaryl, optionally substituted cycloalkyl, or optionally substituted heterocyclyl, wherein the optional substitutions of the aryl, heteroaryl, cycloalkyl, and heterocycloalkyl are CH2OR30, CH2NHR30, CH2NCH3R30, CONR27aR27b, CH2CONR27aR27b, CH2NHCOR26, CH2NCH3COR26, or
特定の実施形態では、ULMは、以下の群から選択される化学構造を有し、

式ULM-c、ULM-d、およびULM-eのR1は、H、エチル、イソプロピル、tert-ブチル、sec-ブチル、シクロプロピル、シクロブチル、シクロペンチル、またはシクロヘキシル、任意に置換されたアルキル、任意に置換されたシクロアルキル、任意に置換されたヒドロキシアルキル、任意に置換されたヘテロアリール、またはハロアルキルであり、
式ULM-c、ULM-d、およびULM-eのR14aは、H、ハロアルキル、任意に置換されたアルキル、メチル、フルオロメチル、ヒドロキシメチル、エチル、イソプロピル、またはシクロプロピルであり、
式ULM-c、ULM-d、およびULM-eのR15は、H、ハロゲン、CN、C≡CH、OH、NO2、任意に置換されたヘテロアリール、任意に置換されたアリール、任意に置換されたアルキル、任意に置換されたハロアルキル、任意に置換されたハロアルコキシ、任意に置換されたシクロアルキル、または任意に置換されたヘテロシクリルからなる群から選択され、
式ULM-c、ULM-d、およびULM-eのXは、C、CH2、またはC=Oであり、
式ULM-c、ULM-d、およびULM-eのR3は、存在しないか、または任意に置換された5または6員ヘテロアリールであり、
破線は、少なくとも1つのPTM、別のULM(ULM’)、または少なくとも1つのPTMもしくはULM’もしくは両方をULMに結合させる化学リンカー部分の結合部位を示す。
In certain embodiments, the ULM has a chemical structure selected from the group consisting of:
R1 in formulae ULM-c, ULM-d, and ULM-e is H, ethyl, isopropyl, tert-butyl, sec-butyl, cyclopropyl, cyclobutyl, cyclopentyl, or cyclohexyl, optionally substituted alkyl, optionally substituted cycloalkyl, optionally substituted hydroxyalkyl, optionally substituted heteroaryl, or haloalkyl;
R14a of formulae ULM-c, ULM-d, and ULM-e is H, haloalkyl, optionally substituted alkyl, methyl, fluoromethyl, hydroxymethyl, ethyl, isopropyl, or cyclopropyl;
R15 of formulae ULM-c, ULM-d, and ULM-e is selected from the group consisting of H, halogen, CN, C≡CH, OH, NO2, optionally substituted heteroaryl, optionally substituted aryl, optionally substituted alkyl, optionally substituted haloalkyl, optionally substituted haloalkoxy, optionally substituted cycloalkyl, or optionally substituted heterocyclyl;
X in formulae ULM-c, ULM-d, and ULM-e is C, CH2, or C=O;
R3 of formulae ULM-c, ULM-d, and ULM-e is absent or an optionally substituted 5- or 6-membered heteroaryl;
The dashed lines indicate the attachment sites of at least one PTM, another ULM (ULM'), or a chemical linker moiety linking at least one PTM or a ULM' or both to a ULM.
特定の実施形態では、ULMは、以下の化学構造に従う基を含み、

式ULM-fのR14aは、H、ハロアルキル、任意に置換されたアルキル、メチル、フルオロメチル、ヒドロキシメチル、エチル、イソプロピル、またはシクロプロピルであり、
式ULM-fのR9は、Hであり、
式ULM-fのR10は、H、エチル、イソプロピル、tert-ブチル、sec-ブチル、シクロプロピル、シクロブチル、シクロペンチル、またはシクロヘキシルであり、
式ULM-fのR11は、

式ULM-fのpは、0、1、2、3、または4であり、
式ULM-fの各R18は、独立して、ハロ、任意に置換されたアルコキシ、シアノ、任意に置換されたアルキル、ハロアルキル、ハロアルコキシまたはリンカーであり、
式ULM-fのR12は、H、C=Oであり、
式ULM-fのR13は、H、任意に置換されたアルキル、任意に置換されたアルキルカルボニル、任意に置換された(シクロアルキル)アルキルカルボニル、任意に置換されたアラルキルカルボニル、任意に置換されたアリールカルボニル、任意に置換された(ヘテロシクリル)カルボニル、または任意に置換されたアラルキルであり、
式ULM-fのR15は、H、ハロゲン、Cl、CN、C≡CH、OH、NO2、任意に置換されたハロアルキル、任意に置換されたヘテロアリール、任意に置換されたアリール、

式中、式ULM-fの破線は、少なくとも1つのPTM、別のULM(ULM’)、または少なくとも1つのPTMもしくはULM’もしくはその両方をULMに結合する化学リンカー部分の結合部位を示す。
In certain embodiments, the ULM comprises a group according to the following chemical structure:
R14a of formula ULM-f is H, haloalkyl, optionally substituted alkyl, methyl, fluoromethyl, hydroxymethyl, ethyl, isopropyl, or cyclopropyl;
R9 of formula ULM-f is H;
R10 of formula ULM-f is H, ethyl, isopropyl, tert-butyl, sec-butyl, cyclopropyl, cyclobutyl, cyclopentyl, or cyclohexyl;
R11 in the formula ULM-f is
In the formula ULM-f, p is 0, 1, 2, 3, or 4;
Each R of formula ULM-f is independently halo, optionally substituted alkoxy, cyano, optionally substituted alkyl, haloalkyl, haloalkoxy, or a linker;
R12 of formula ULM-f is H, C=O;
R13 of formula ULM-f is H, optionally substituted alkyl, optionally substituted alkylcarbonyl, optionally substituted (cycloalkyl)alkylcarbonyl, optionally substituted aralkylcarbonyl, optionally substituted arylcarbonyl, optionally substituted (heterocyclyl)carbonyl, or optionally substituted aralkyl;
R15 of formula ULM-f is H, halogen, Cl, CN, C≡CH, OH, NO2, optionally substituted haloalkyl, optionally substituted heteroaryl, optionally substituted aryl,
wherein the dashed lines in formula ULM-f indicate the site of attachment of at least one PTM, another ULM (ULM'), or a chemical linker moiety that connects at least one PTM or ULM' or both to a ULM.
特定の実施形態では、ULMは、以下の構造から選択される:

特定の実施形態では、ULMは、以下の構造から選択される:


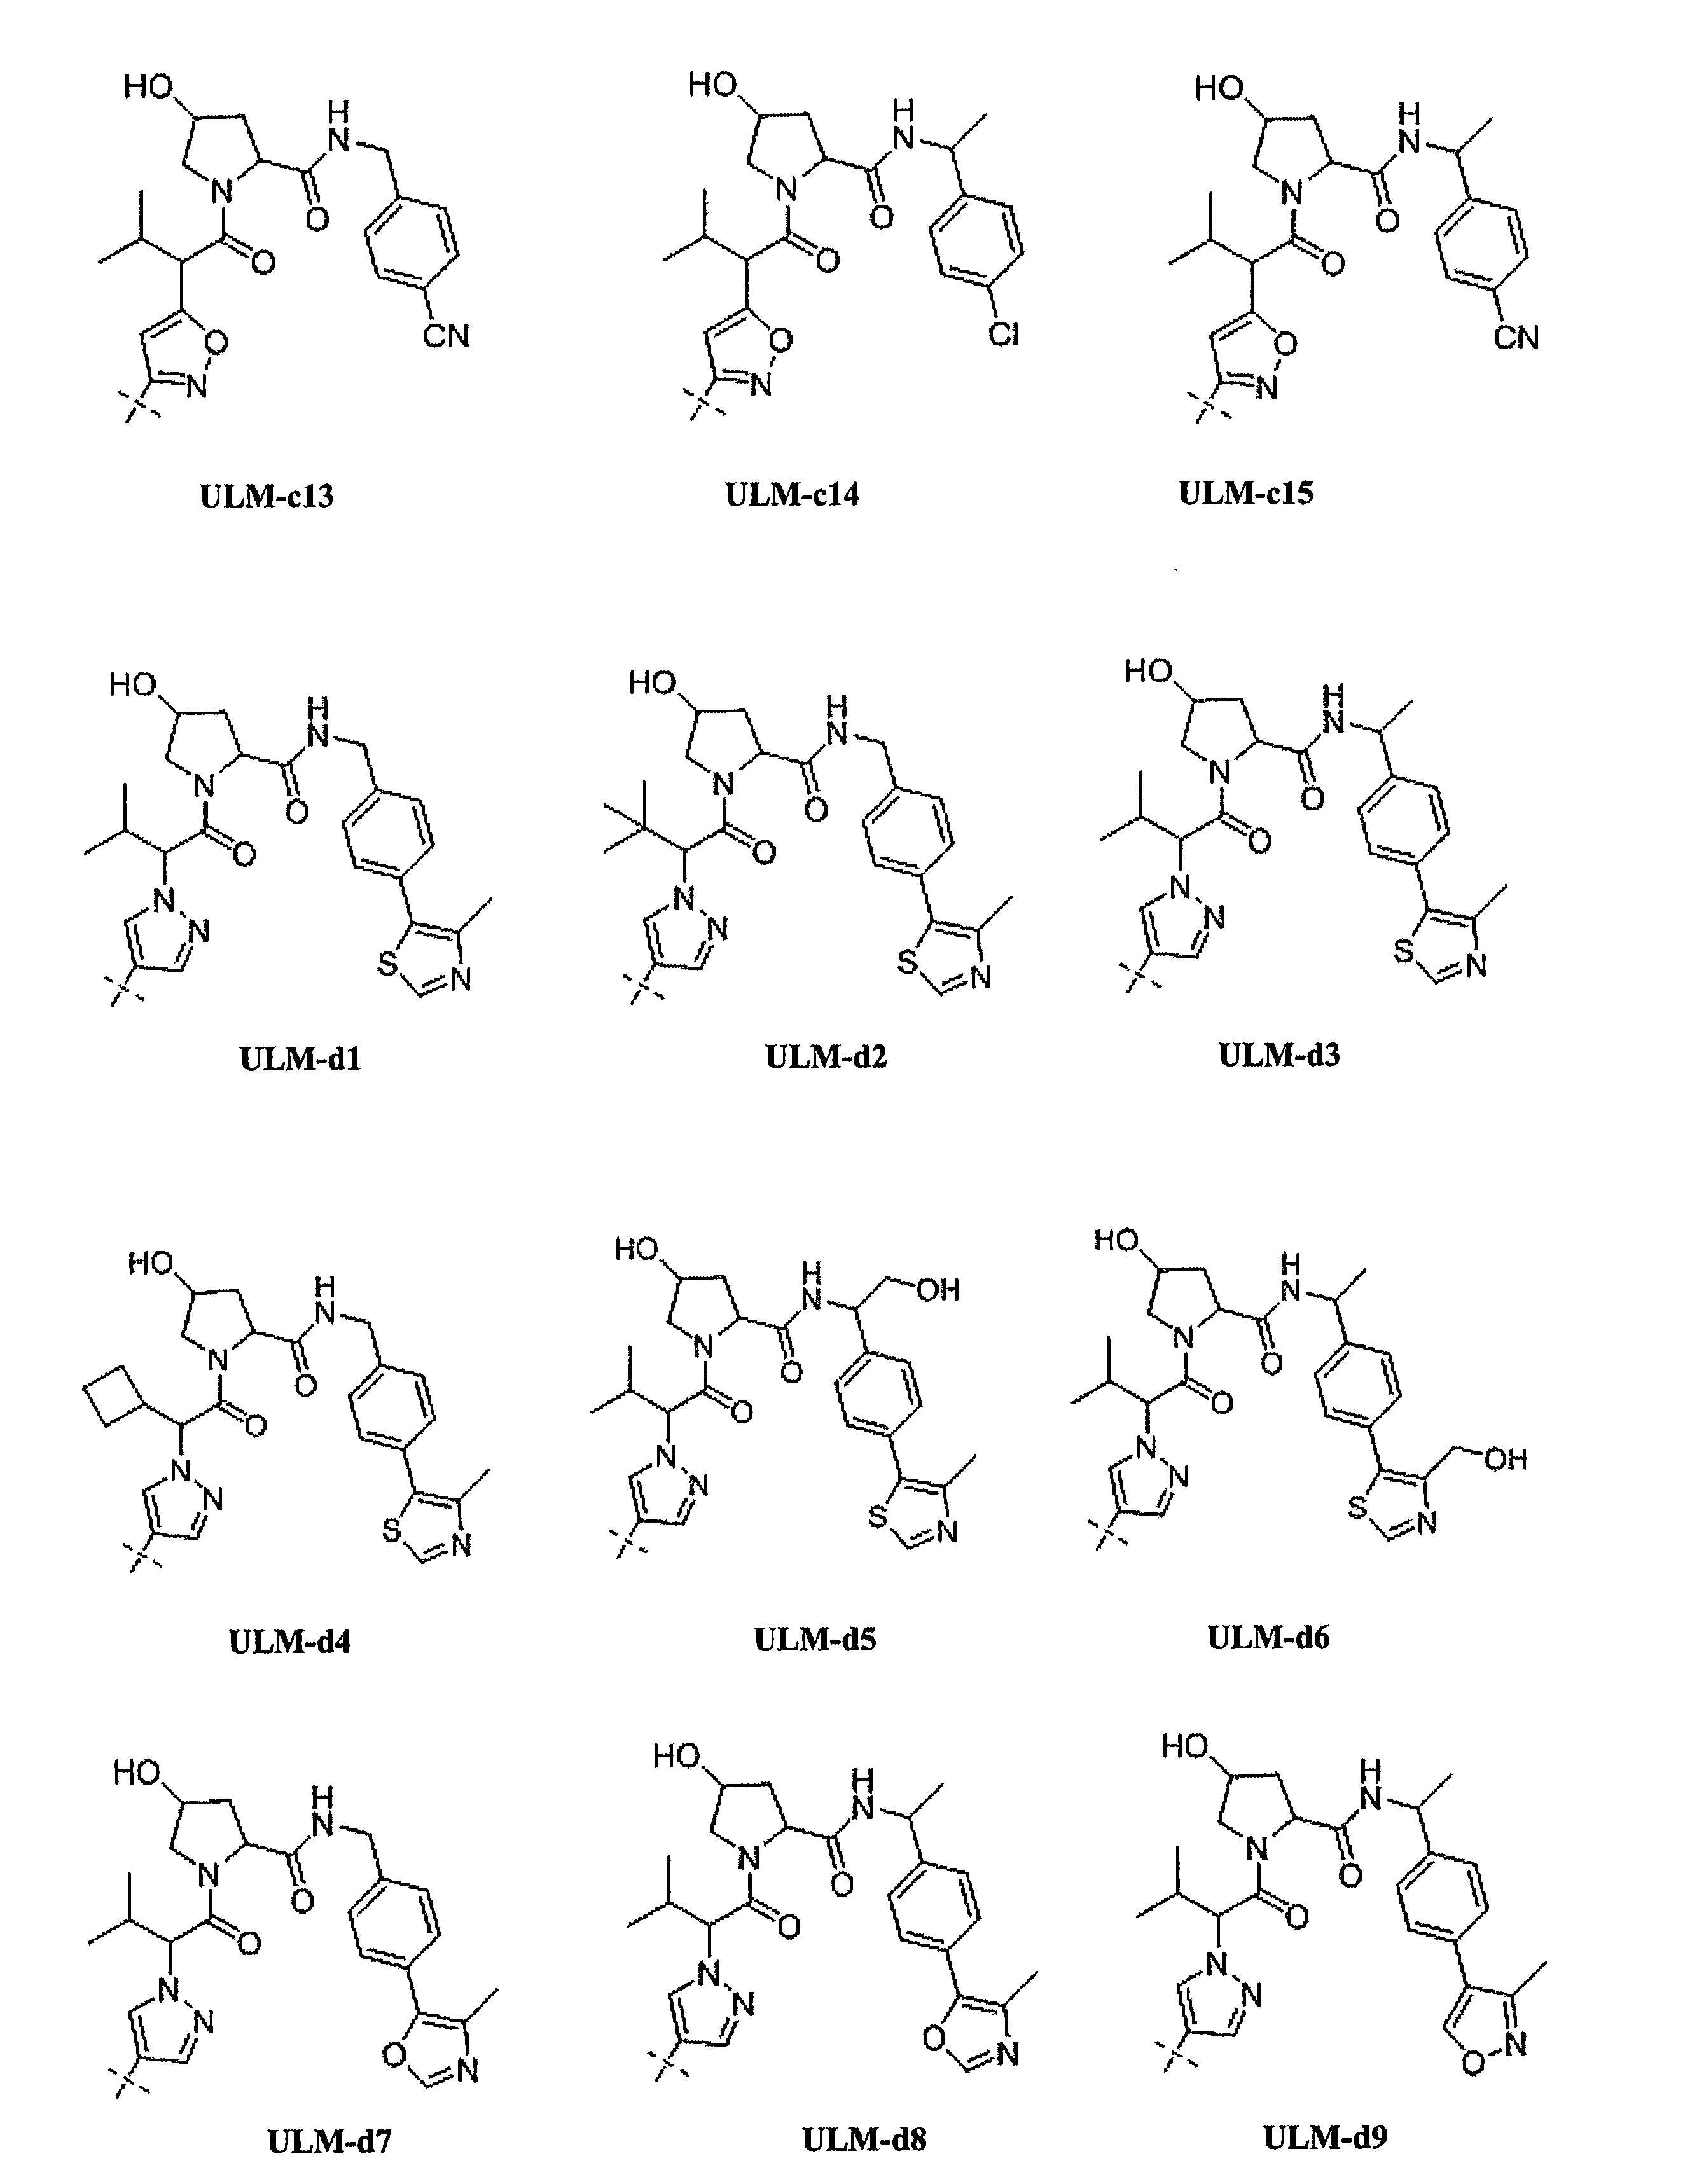
1つの実施形態では、ULM-a1からULM-a15、ULM-b1からULM-b12、ULM-c1からULM-c15、およびULM-d1からULM-d9のフェニル環はエステルとして官能基化され、それをプロドラッグの一部とすることができる。 In one embodiment, the phenyl rings of ULM-a1 through ULM-a15, ULM-b1 through ULM-b12, ULM-c1 through ULM-c15, and ULM-d1 through ULM-d9 are functionalized as esters, which can be part of a prodrug.
特定の実施形態では、ULM-a1からULM-a15、ULM-b1からULM-b12、ULM-c1からULM-c15、およびULM-d1からULM-d9のピロリジン環上のヒドロキシル基はそれぞれエステル結合されたプロドラッグ部分を含む。 In certain embodiments, the hydroxyl group on the pyrrolidine ring of ULM-a1 through ULM-a15, ULM-b1 through ULM-b12, ULM-c1 through ULM-c15, and ULM-d1 through ULM-d9 each contain an ester-linked prodrug moiety.
本明細書に記載される態様または実施形態のいずれかにおいて、ULM、および存在する場合にはULM’、またはその薬学的に許容可能な塩、立体異性体、溶媒和物もしくは多形体は、それぞれ独立して、以下の化学構造に従う基である:

ULM-gのR1’は、任意に置換されたC1-C6アルキル基、任意に置換された-(CH2)nOH、任意に置換された-(CH2)nSH、任意に置換された(CH2)n-O-(C1-C6)アルキル基、エポキシド部分WCOCW(式中、各Wは、独立して、HまたはC1-C3アルキル基である)を含有する任意に置換された(CH2)n-WCOCW-(C0-C6)アルキル基、任意に置換された-(CH2)nCOOH、任意に置換された-(CH2)nC(O)-(C1-C6アルキル)、任意に置換された-(CH2)nNHC(O)-R″、任意に置換された-(CH2)nC(O)-N(R″)2、任意に置換された-(CH2)nOC(O)-N(R″)2、-(CH2O)nH、任意に置換された-(CH2)nOC(O)-(C1-C6アルキル)、任意に置換された-(CH2)nC(O)-O-(C1-C6アルキル)、任意に置換された-(CH2O)nCOOH、任意に置換された-(OCH2)nO-(C1-C6アルキル)、任意に置換された-(CH2O)nC(O)-(C1-C6アルキル)、任意に置換された-(OCH2)nNHC(O)-R″、任意に置換された-(CH2O)nC(O)-N(R″)2、-(CH2CH2O)nH、任意に置換された-(CH2CH2O)nCOOH、任意に置換された-(OCH2CH2)nO-(C1-C6アルキル)、任意に置換された-(CH2CH2O)nC(O)-(C1-C6アルキル)、任意に置換された-(OCH2CH2)nNHC(O)-R″、任意に置換された-(CH2CH2O)nC(O)-N(R″)2、任意に置換された-SO2RS、任意に置換されたS(O)RS、NO2、CN、またはハロゲン(F、Cl、Br、I、好ましくは、FまたはCl)であり、
ULM-gの各R″は、独立して、H、または1個もしくは2個のヒドロキシル基または最大3個のハロゲン基(好ましくは、フッ素)で任意に置換され得るC1-C6アルキル基であり、
ULM-gのRSが、C1-C6アルキル基、任意に置換されたアリール、ヘテロアリール、もしくはヘテロシクリル基、または-(CH2)mN(R″)2基であり、
ULM-gのXおよびX’は、各々独立して、C=O、C=S、-S(O)、S(O)2であり、(好ましくは、XおよびX’は、両方ともC=Oであり)、
ULM-gのR2’は、任意に置換された-(CH2)n-(C=O)u(NR″)v(SO2)wアルキリ基、任意に置換された-(CH2)n-(C=O)u(NR″)v(SO2)wNR1NR2N基、任意に置換された-(CH2)n-(C=O)u(NR″)v(SO2)w-アリール、任意に置換された-(CH2)n-(C=O)u(NR″)v(SO2)w-ヘテロアリール、任意に置換された-(CH2)n-(C=O)vNR″(SO2)w-ヘテロシクリル、任意に置換された-NR″-(CH2)n-C(O)u(NR″)v(SO2)w-アルキル、任意に置換された-NR″-(CH2)n-C(O)u(NR″)v(SO2)w-NR1NR2N、任意に置換された-NR″-(CH2)n-C(O)u(NR″)v(SO2)w-NR″C(O)R1N、任意に置換された-NR″-(CH2)n-(C=O)u(NR″)v(SO2)w-アリール、任意に置換された-NR″-(CH2)n-(C=O)u(NR″)v(SO2)w-ヘテロアリール、もしくは任意に置換された-NR″-(CH2)n-(C=O)vNR″(SO2)w-ヘテロシクリル、任意に置換された-XR2’-アルキリ基、任意に置換された-XR2’-アリール基、任意に置換された-XR2’-ヘテロアリール基、任意に置換された-XR2’-ヘテロシクリル基であり、
ULM-gのR3’は、任意に置換されたアルキル、任意に置換された-(CH2)n-(O)u(NR″)v(SO2)w-アルキル、任意に置換された-(CH2)n-C(O)u(NR″)v(SO2)w-NR1NR2N、任意に置換された-(CH2)n-C(O)u(NR″)v(SO2)w-NR″C(O)R1N、任意に置換された-(CH2)n-C(O)u(NR″)v(SO2)w-C(O)(R″)2、任意に置換された-(CH2)n-C(O)u(NR″)v(SO2)w-アリール、任意に置換された-(CH2)n-C(O)u(NR″)v(SO2)w-ヘテロアリール、任意に置換された-(CH2)n-C(O)u(NR″)v(SO2)w-ヘテロシクリル、任意に置換された-NR″-(CH2)n-C(O)u(NR″)v(SO2)w-アルキル、任意に置換された-NR″-(CH2)n-C(O)u(NR″)v(SO2)w-NR1NR2N、任意に置換された-NR″-(CH2)n-C(O)u(NR″)v(SO2)w-NR″C(O)R1N、任意に置換された-NR″-(CH2)n-C(O)u(NR″)v(SO2)w-アリール、任意に置換された-NR″-(CH2)n-C(O)u(NR″)v(SO2)w-ヘテロアリール、任意に置換された-NR1-(CH2)n-C(O)u(NR″)v(SO2)w-ヘテロシクリル、任意に置換された-O-(CH2)n-(C=O)u(NR″)v(SO2)w-アルキル、任意に置換された-O-(CH2)n-(C=O)u(NR″)v(SO2)w-NR1NR2N、任意に置換された-O-(CH2)n-(C=O)u(NR″)v(SO2)w-NR″C(O)R1N、任意に置換された-O-(CH2)n-(C=O)u(NR″)v(SO2)w-アリール、任意に置換された-O-(CH2)n-(C=O)u(NR″)v(SO2)w-ヘテロアリール、もしくは任意に置換された-O-(CH2)n-(C=O)u(NR″)v(SO2)w-ヘテロシクリル、-(CH2)n-(V)n’-(CH2)n-(V)n’-アルキル基、任意に置換された-(CH2)n-(V)n’-(CH2)n-(V)n’-アリール基、任意に置換された-(CH2)n-(V)n’-(CH2)n-(V)n’-ヘテロアリール基、任意に置換された-(CH2)n-(V)n’-(CH2)n-(V)n’-ヘテロシクリル’基、任意に置換された-(CH2)n-N(R1’)(C=O)m’-(V)n’-アルキル基、任意に置換された-(CH2)n-N(R1’)(C=O)m’-(V)n’-アリール基、任意に置換された-(CH2)n-N(R1’)(C=O)m’-(V)n’-ヘテロアリール基、任意に置換された-(CH2)n-N(R1’)(C=O)m’-(V)n’-ヘテロシクリル基、任意に置換された-XR3’-アルキル基、任意に置換された-XR3’-アリール基、任意に置換された-XR3’-ヘテロアリール基、任意に置換された-XR3’-ヘテロシクリル基であり、
ULM-gのR1NおよびR2Nは、各々独立して、H、1個もしくは2個のヒドロキシル基および最大3個のハロゲン基で任意に置換されたC1-C6アルキル、または任意に置換された-(CH2)n-アリール、-(CH2)n-ヘテロアリール、もしくは-(CH2)n-ヘテロシクリル基であり、
ULM-gのVは、O、SまたはNR1であり、
ULM-gの各R1’は、独立して、H、またはC1-C3アルキル基であり、
ULM-gのXR2’およびXR3’は、各々独立して、任意に置換された-CH2)n-、-CH2)n-CH(Xv)=CH(Xv)-(cisまたはtrans)、-CH2)n-CH≡CH-、-(CH2CH2O)n-、またはC3-C6シクロアルキル基であり、式中、Xvは、H、ハロ、または任意に置換されたC1-C3アルキル基であり、
ULM-gの各mは独立して、0、1、2、3、4、5、6であり、
ULM-gの各m’は独立して、0または1であり、
ULM-gの各nは独立して、0、1、2、3、4、5、6であり、
ULM-gの各n’は独立して、0または1であり、
ULM-gの各uは独立して、0または1であり、
ULM-gの各vは独立して、0または1であり、
ULM-gの各wは独立して、0または1であり、および
ULM-gのR1’、R2’、R3’、XおよびX’のうちのいずれか1つ以上が任意で改変されて、PTMがULM’ではない場合はリンカー基を介してPTM基に共有結合され、PTMがULM’である場合は、ULMおよびULM’の各々のR1’、R2’、R3’、XならびにX’のうちのいずれか1つ以上が任意で改変されて、直接またはリンカー基を介して互いに共有結合される。
In any of the aspects or embodiments described herein, ULM, and, if present, ULM′, or a pharma- ceutically acceptable salt, stereoisomer, solvate or polymorph thereof, are each independently a group conforming to the following chemical structure:
R1' of ULM-g is an optionally substituted C1-C6 alkyl group, an optionally substituted -(CH2)nOH, an optionally substituted -(CH2)nSH, an optionally substituted (CH2)n-O-(C1-C6)alkyl group, an optionally substituted (CH2)n-WCOCW-(C0-C6)alkyl group containing an epoxide moiety WCOCW (where each W is independently H or a C1-C3 alkyl group), an optionally substituted -(CH 2) nCOOH, optionally substituted -(CH2)nC(O)-(C1-C6 alkyl), optionally substituted -(CH2)nNHC(O)-R", optionally substituted -(CH2)nC(O)-N(R")2, optionally substituted -(CH2)nOC(O)-N(R")2, -(CH2O)nH, optionally substituted -(CH2)nOC(O)-(C1-C6 alkyl), optionally substituted -(CH2)nC(O)-O-(C1-C6 optionally substituted -(CH2O)nCOOH, optionally substituted -(OCH2)nO-(C1-C6 alkyl), optionally substituted -(CHO)nC(O)-(C1-C6 alkyl), optionally substituted -(OCH2)nNHC(O)-R", optionally substituted -(CHO)nC(O)-N(R")2, -(CH2CH2O)nH, optionally substituted -(CH2CH2O)nCOOH, optionally substituted -( optionally substituted —(CHCHO)C(O)—N(R″)2, optionally substituted —SORS, optionally substituted S(O)RS, NO, CN, or halogen (F, Cl, Br, I, preferably F or Cl);
Each R″ in ULM-g is independently H or a C1-C6 alkyl group which may be optionally substituted with one or two hydroxyl groups or up to three halogen groups (preferably fluorine);
RS of ULM-g is a C1-C6 alkyl group, an optionally substituted aryl, heteroaryl, or heterocyclyl group, or a -(CH2)mN(R'')2 group;
X and X' of ULM-g are each independently C=O, C=S, -S(O), S(O)2 (preferably, X and X' are both C=O);
R2' of ULM-g is an optionally substituted -(CH2)n-(C=O)u(NR'')v(SO2)w alkyl group, an optionally substituted -(CH2)n-(C=O)u(NR'')v(SO2)w NR1NR2N group, an optionally substituted -(CH2)n-(C=O)u(NR'')v(SO2)w aryl, an optionally substituted -(CH2)n-(C=O)u(NR'')v(SO2)w-heteroaryl, an optionally substituted -(CH2)n-(C=O)vNR''(SO2)w-heterocyclyl, an optionally substituted -NR''-(CH2)n-C(O)u(NR'')v(SO2)w alkyl, an optionally substituted -NR''-(CH2)n-C(O)u(NR'')v (SO2)w-NR1NR2N, optionally substituted -NR"-(CH2)n-C(O)u(NR")v(SO2)w-NR"C(O)RN, optionally substituted -NR"-(CH2)n-(C=O)u(NR")v(SO2)w-aryl, optionally substituted -NR"-(CH2)n-(C=O)u(NR")v(SO2)w-heteroaryl or optionally substituted -NR"-(CH2)n-(C=O)vNR"(SO2)w-heterocyclyl, optionally substituted -XR2'-alkyl group, optionally substituted -XR2'-aryl group, optionally substituted -XR2'-heteroaryl group, or optionally substituted -XR2'-heterocyclyl group;
R3' of ULM-g is optionally substituted alkyl, optionally substituted -(CH2)n-(O)u(NR'')v(SO2)w-alkyl, optionally substituted -(CH2)n-C(O)u(NR'')v(SO2)w-NR1NR2N, optionally substituted -(CH2)n-C(O)u(NR'')v(SO2)w-NR''C(O)R1N, optionally substituted -(C H2)n-C(O)u(NR'')v(SO2)w-C(O)(R'')2, optionally substituted -(CH2)n-C(O)u(NR'')v(SO2)w-aryl, optionally substituted -(CH2)n-C(O)u(NR'')v(SO2)w-heteroaryl, optionally substituted -(CH2)n-C(O)u(NR'')v(SO2)w-heterocyclyl, optionally substituted optionally substituted -NR''-(CH2)n-C(O)u(NR'')v(SO2)w-alkyl, optionally substituted -NR''-(CH2)n-C(O)u(NR'')v(SO2)w-NR1NR2N, optionally substituted -NR''-(CH2)n-C(O)u(NR'')v(SO2)w-NR''C(O)R1N, optionally substituted -NR''-(CH2)n-C(O)u(NR'')v(SO2)w-NR''C(O)R1N optionally substituted —NR″-(CH2)n-C(O)u(NR″)v(SO2)w-aryl, optionally substituted —NR″-(CH2)n-C(O)u(NR″)v(SO2)w-heteroaryl, optionally substituted —NR1-(CH2)n-C(O)u(NR″)v(SO2)w-heterocyclyl, optionally substituted —O—(CH2)n-(C═O)u(NR″)v(SO2)w-alkyl, optionally substituted —O—( CH2)n-(C=O)u(NR'')v(SO2)w-NR1NR2N, optionally substituted -O-(CH2)n-(C=O)u(NR'')v(SO2)w-NR''C(O)R1N, optionally substituted -O-(CH2)n-(C=O)u(NR'')v(SO2)w-aryl, optionally substituted -O-(CH2)n-(C=O)u(NR'')v(SO2)w- Heteroaryl, or optionally substituted -O-(CH2)n-(C=O)u(NR'')v(SO2)w-heterocyclyl, -(CH2)n-(V)n'-(CH2)n-(V)n'-alkyl group, optionally substituted -(CH2)n-(V)n'-(CH2)n-(V)n'-aryl group, optionally substituted -(CH2)n-(V)n'-(CH2)n-( V) n'-heteroaryl groups, optionally substituted -(CH2)n-(V) n'-(CH2)n-(V) n'-heterocyclyl' groups, optionally substituted -(CH2)n-N(R1')(C=O)m'-(V) n'-alkyl groups, optionally substituted -(CH2)n-N(R1')(C=O)m'-(V) n'-aryl groups, optionally substituted -(CH2)n-N(R1')(C=O)m'-(V) n'-heteroaryl groups, optionally substituted -(CH2)n-N(R1')(C=O)m'-(V) n'-heterocyclyl groups, optionally substituted -XR3'-alkyl groups, optionally substituted -XR3'-aryl groups, optionally substituted -XR3'-heteroaryl groups, optionally substituted -XR3'-heterocyclyl groups;
R1N and R2N of ULM-g are each independently H, C1-C6 alkyl optionally substituted with one or two hydroxyl groups and up to three halogen groups, or an optionally substituted -(CH2)n-aryl, -(CH2)n-heteroaryl, or -(CH2)n-heterocyclyl group;
V of ULM-g is O, S or NR1;
Each R1' in ULM-g is independently H or a C1-C3 alkyl group;
XR2' and XR3' of ULM-g are each independently an optionally substituted -CH2)n-, -CH2)n-CH(Xv)=CH(Xv)-(cis or trans), -CH2)n-CH≡CH-, -(CH2CHO)n-, or a C3-C6 cycloalkyl group, where Xv is H, halo, or an optionally substituted C1-C3 alkyl group;
Each m in ULM-g is independently 0, 1, 2, 3, 4, 5, or 6;
Each m' in ULM-g is independently 0 or 1;
Each n in ULM-g is independently 0, 1, 2, 3, 4, 5, or 6;
Each n' in ULM-g is independently 0 or 1;
Each u in ULM-g is independently 0 or 1;
Each v in ULM-g is independently 0 or 1;
each w of ULM-g is independently 0 or 1; and any one or more of R1', R2', R3', X and X' of ULM-g are optionally modified to be covalently bonded to the PTM group via a linker group if the PTM is not a ULM', and any one or more of R1', R2', R3', X and X' of each of ULM and ULM' are optionally modified to be covalently bonded to each other directly or via a linker group if the PTM is a ULM'.
本明細書に記載される態様または実施形態のいずれかにおいて、ULM、および存在する場合にはULM’、またはその薬学的に許容可能な塩、鏡像異性体、ジアステレオマー、溶媒和物もしくは多形体は、それぞれ独立して、以下の化学構造に従う基である:

ULM-hの各R1’、R2’、およびR3’は、上記と同じであり、Xが、C=O、C=S、-S(O)基、またはS(O)2基、より好ましくは、C=O基であり、
ULM-hのR1’、R2’、およびR3’のうちのいずれか1つ以上が任意で改変されて、PTMがULM’ではない場合は、PTM基にさらに共有結合されるリンカー基を結合し、またはPTMがULM’である場合は、ULMおよびULM’の各々のR1’、R2’、R3’のうちのいずれか1つ以上が任意で改変されて、直接、またはリンカー基を介して互いに共有結合される。
In any of the aspects or embodiments described herein, ULM, and, if present, ULM′, or a pharma- ceutically acceptable salt, enantiomer, diastereomer, solvate or polymorph thereof, are each independently a group conforming to the following chemical structure:
each R1', R2', and R3' of ULM-h is the same as above, and X is C=O, C=S, -S(O) group, or S(O)2 group, more preferably C=O group;
Any one or more of R1', R2', and R3' of ULM-h are optionally modified to attach a linker group that is further covalently attached to the PTM group if the PTM is not a ULM', or any one or more of R1', R2', R3' of each of the ULM and ULM' are optionally modified to be covalently attached to each other directly or via a linker group if the PTM is a ULM'.
本明細書に記載される態様または実施形態のいずれかにおいて、ULM、および存在する場合にはULM’、またはその薬学的に許容可能な塩、鏡像異性体、ジアステレオマー、溶媒和物もしくは多形体は、それぞれ独立して、以下の化学構造に従う:
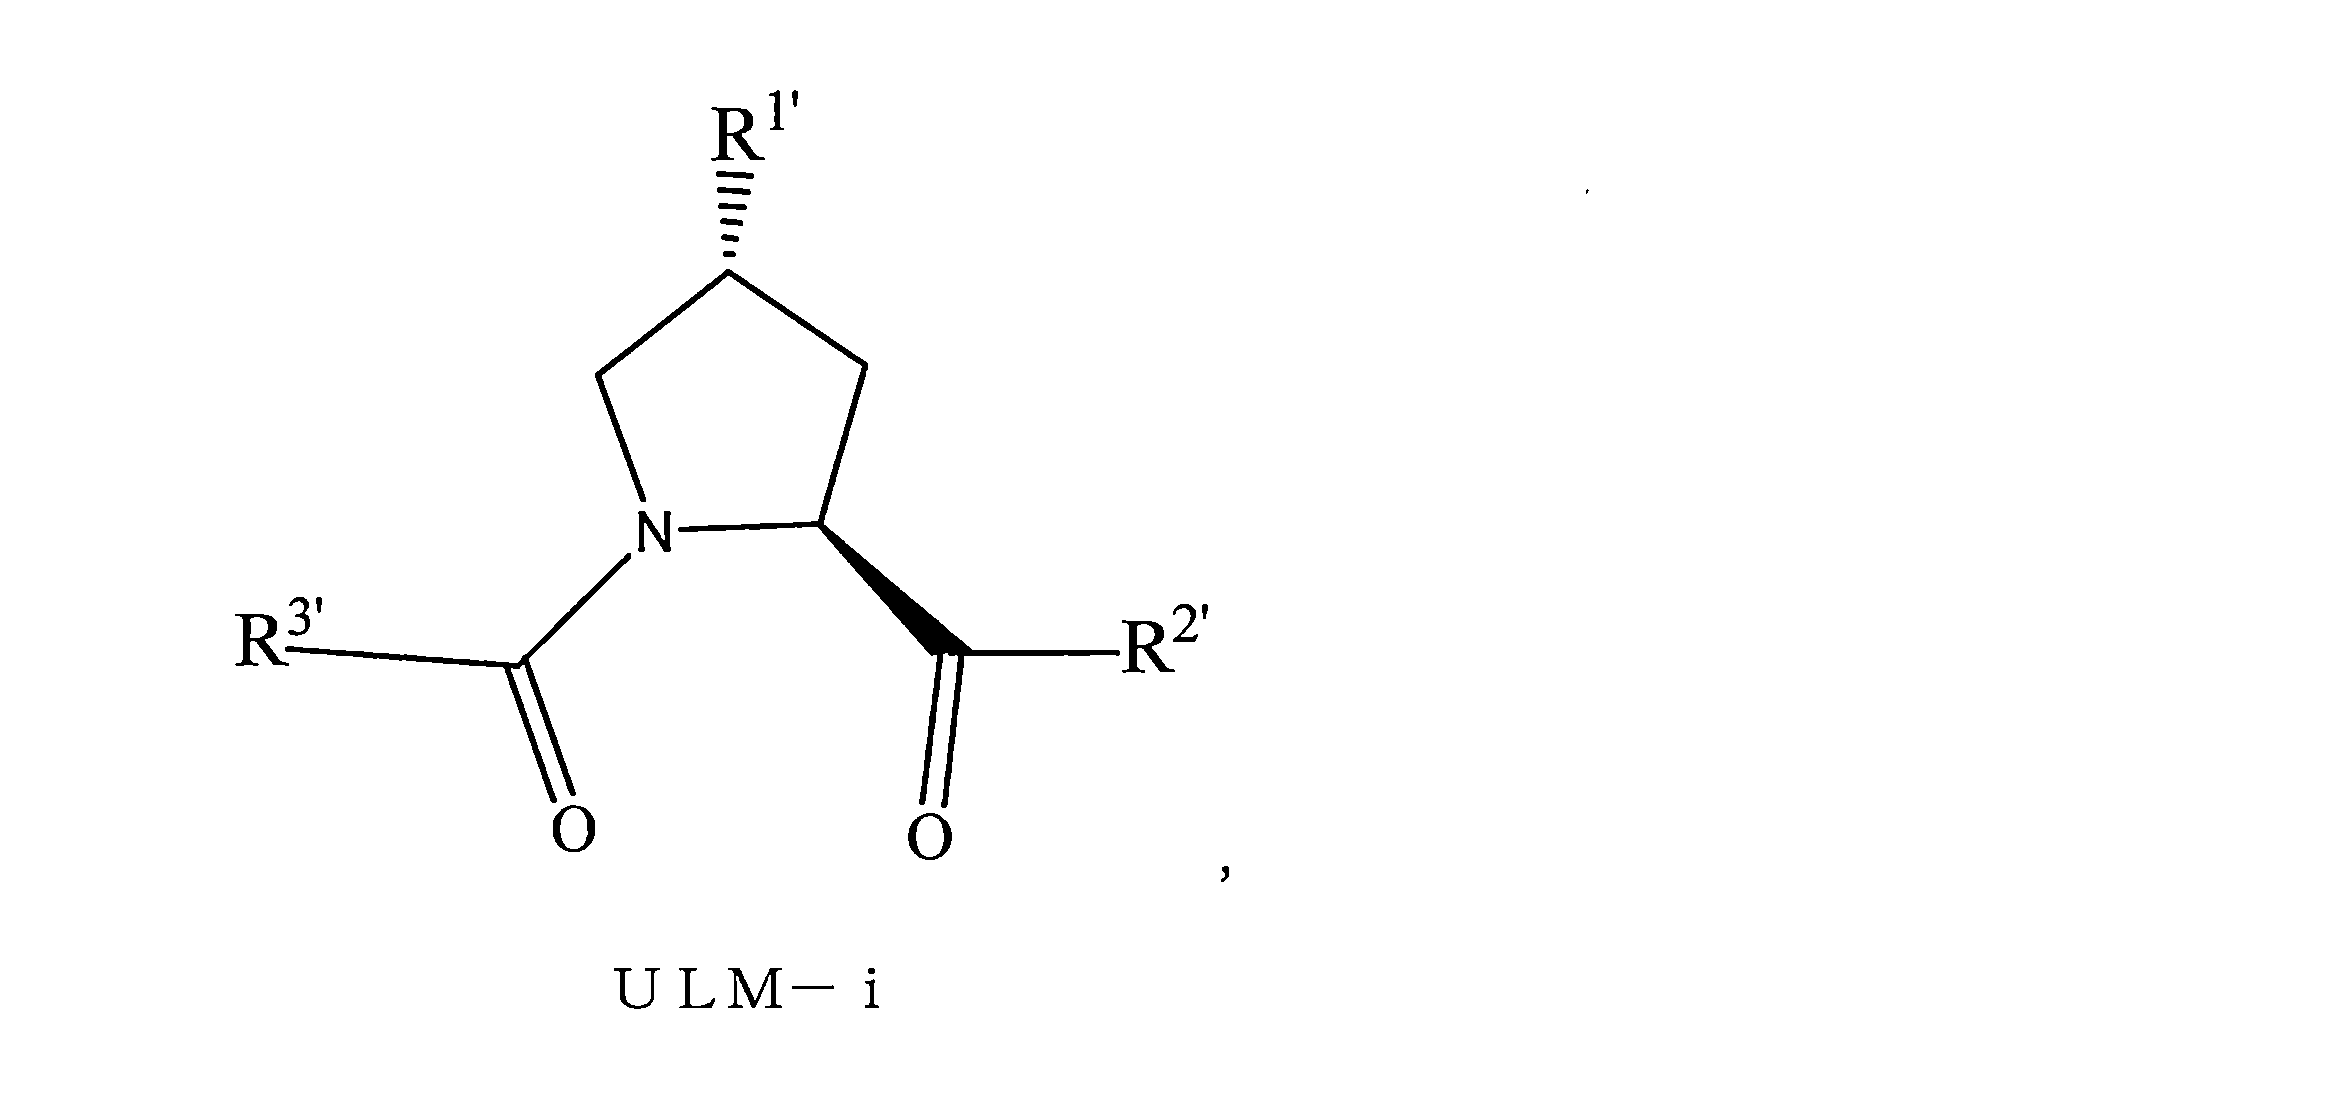
ULM-IのR1’、R2’、およびR3’のうちのいずれか1つ以上が任意で改変されて、PTMがULM’ではない場合は、PTM基にさらに共有結合されるリンカー基を結合し、またはPTMがULM’である場合は、ULMおよびULM’の各々のR1’、R2’、R3’のうちのいずれか1つ以上が任意で改変されて、直接、またはリンカー基を介して互いに共有結合される。
In any of the aspects or embodiments described herein, ULM, and, if present, ULM′, or a pharma- ceutically acceptable salt, enantiomer, diastereomer, solvate or polymorph thereof, each independently conforms to the following chemical structure:
Any one or more of R1', R2', and R3' of ULM-I are optionally modified to attach a linker group that is further covalently attached to the PTM group if the PTM is not a ULM', or any one or more of R1', R2', R3' of each of ULM and ULM' are optionally modified to be covalently attached to each other directly or via a linker group if the PTM is a ULM'.
本開示のさらなる好ましい態様では、ULM-g~ULM-iのR1’は、好ましくは、ヒドロキシル基、または化合物が活性化合物のプロドラッグ形態を表すようにヒドロキシルもしくはカルボン酸基に代謝され得る基である。好ましいR1’基の例としては例えば、-(CH2)nOH、(CH2)n-O-(C1-C6)アルキル基、-(CH2)nCOOH、-(CH2O)nH、任意に置換された-(CH2)nOC(O)-(C1-C6アルキル)、または任意に置換された-(CH2)nC(O)-O-(C1-C6アルキル)が挙げられ、式中、nは0または1である。式中、R1’は、カルボン酸基、ヒドロキシル基もしくはアミン基であるか、またはそれを含有し、当該ヒドロキシル基、カルボン酸基またはアミン(それら各々が任意で置換され得る)はさらに化学改変されて、PTM基(ULM’基を含む)に結合されるリンカー基に対する共有結合を提供し得る。 In further preferred aspects of the present disclosure, R1' in ULM-g to ULM-i is preferably a hydroxyl group or a group that can be metabolized to a hydroxyl or carboxylic acid group such that the compound represents a prodrug form of the active compound. Examples of preferred R1' groups include, for example, -(CH2)nOH, (CH2)n-O-(C1-C6) alkyl groups, -(CH2)nCOOH, -(CHO)nH, optionally substituted -(CH2)nOC(O)-(C1-C6 alkyl), or optionally substituted -(CH2)nC(O)-O-(C1-C6 alkyl), where n is 0 or 1. Wherein R1' is or contains a carboxylic acid group, a hydroxyl group, or an amine group, each of which may be optionally substituted, which may be further chemically modified to provide a covalent bond to a linker group that is attached to the PTM group (including the ULM' group).
ULM-gおよびULM-hのXおよびX’は、存在する場合、好ましくはC=O、C=S、-S(O)基またはS(O)2基であり、より好ましくはC=O基である。 X and X' in ULM-g and ULM-h, when present, are preferably C=O, C=S, -S(O) or S(O)2 groups, more preferably C=O groups.
ULM-g~ULM-iのR2’は、好ましくは、任意に置換された-NH-T-アリール、任意に置換された-N(CH3)-T-アリール、任意に置換された-NH-T-ヘテロアリール基、任意に置換された-N(CH3)-T-ヘテロアリール、任意に置換された-NH-T-ヘテロシクリル、または任意に置換された-N(CH3)-T-ヘテロシクリル、好ましくはHであり、Tは、任意に置換された-(CH2)n-基であり、メチレン基の各々は、1個もしくは2個の置換基で任意に置換され得、好ましくは、ハロゲン、本明細書に別途記載されるアミノ酸側鎖、またはC1-C3アルカリ基、好ましくは任意に置換され得る1個もしくは2個のメチル基から選択され、nは、0~6、多くの場合0、1、2、または3、好ましくは0または1である。あるいは、Tはまた、-(CH2O)n-基、-(OCH2)n-基、-(CH2CH2O)n-基、-(OCH2CH2)n-基であり得、それらのすべての基は任意に置換されている。 R2' in ULM-g to ULM-i is preferably an optionally substituted -NH-T-aryl, an optionally substituted -N(CH3)-T-aryl, an optionally substituted -NH-T-heteroaryl group, an optionally substituted -N(CH3)-T-heteroaryl, an optionally substituted -NH-T-heterocyclyl, or an optionally substituted -N(CH3)-T-heterocyclyl, preferably H; T is an optionally substituted -(CH2)n- group, each of the methylene groups may be optionally substituted with one or two substituents, preferably selected from halogen, an amino acid side chain as described elsewhere herein, or a C1-C3 alkyl group, preferably one or two methyl groups which may be optionally substituted; and n is 0 to 6, often 0, 1, 2, or 3, preferably 0 or 1. Alternatively, T can also be a -(CH2O)n- group, a -(OCH2)n- group, a -(CH2CH2O)n- group, or a -(OCH2CH2)n- group, all of which are optionally substituted.
ULM-g~ULM-iのR2’のための好ましいアリール基としては、任意に置換されたフェニル基またはナフチル基、好ましくはフェニル基が挙げられ、式中、フェニルまたはナフチル基は、リンカー基を有するPTM(ULM’基を含む)に連結され、および/または任意にハロゲン(好ましくはFまたはCl)、アミン、モノアルキル、もしくはジアルキルアミン(好ましくはジメチルアミン)、F、Cl、OH、COOH、C1-C6アルキル、好ましくはCH3、CF3、OMe、OCF3、NO2、またはCN基(それらの各々が、フェニル環のオーソ、メタ、および/またはパラ位置、好ましくはパラ位置で置換され得る)で任意に置換され、任意に置換されたフェニル基(フェニル基自体が、リンカー基を有するULM’を含むPTM基に任意に連結されている)、および/またはF、Cl、OH、COOH、CH3、CF3、OMe、OCF3、NO2、もしくはCN基のうちの少なくとも1つで(フェニル環のオーソ、メタ、および/またはパラ位置、好ましくはパラ位置で)置換され、任意に置換され得るナフチル基、任意に置換されたヘテロアリール、好ましくはメチル置換イソキサゾールを含む任意に置換されたイソキサゾール、メチル置換オキサゾールを含む任意に置換されたオキサゾール、メチル置換チアゾールを含む任意に置換されたチアゾール、メチル置換イソチアゾールを含む任意に置換されたイソチアゾール、メチル置換ピロールを含む任意に置換されたピロール、メチルイミダゾールを含む任意に置換されたイミダゾール、任意に置換されたベンズイミダゾールまたはメトキシベンジルイミダゾール、任意に置換されたオキシイミダゾールまたはメチルオキシイミダゾール、メチルジアゾール基を含む任意に置換された置換ジアゾール基、メチル置換トリアゾール基を含む任意に置換されたトリアゾール基、ハロ(好ましくは、F)を含む任意に置換されたピリジン基、もしくはメチル置換ピリジン基、またはオキサピリジン基(ピリジン基が酸素によってフェニル基に結合されている場合)、任意に置換されたフラン、任意に置換されたベンゾフラン、任意に置換されたジヒドロベンゾフラン、任意に置換されたインドール、インドリジン、またはアザインドリジン(2、3、または4-アザインドリジン)、任意に置換されたキノリン、以下の化学構造に従う任意に置換された基であり、
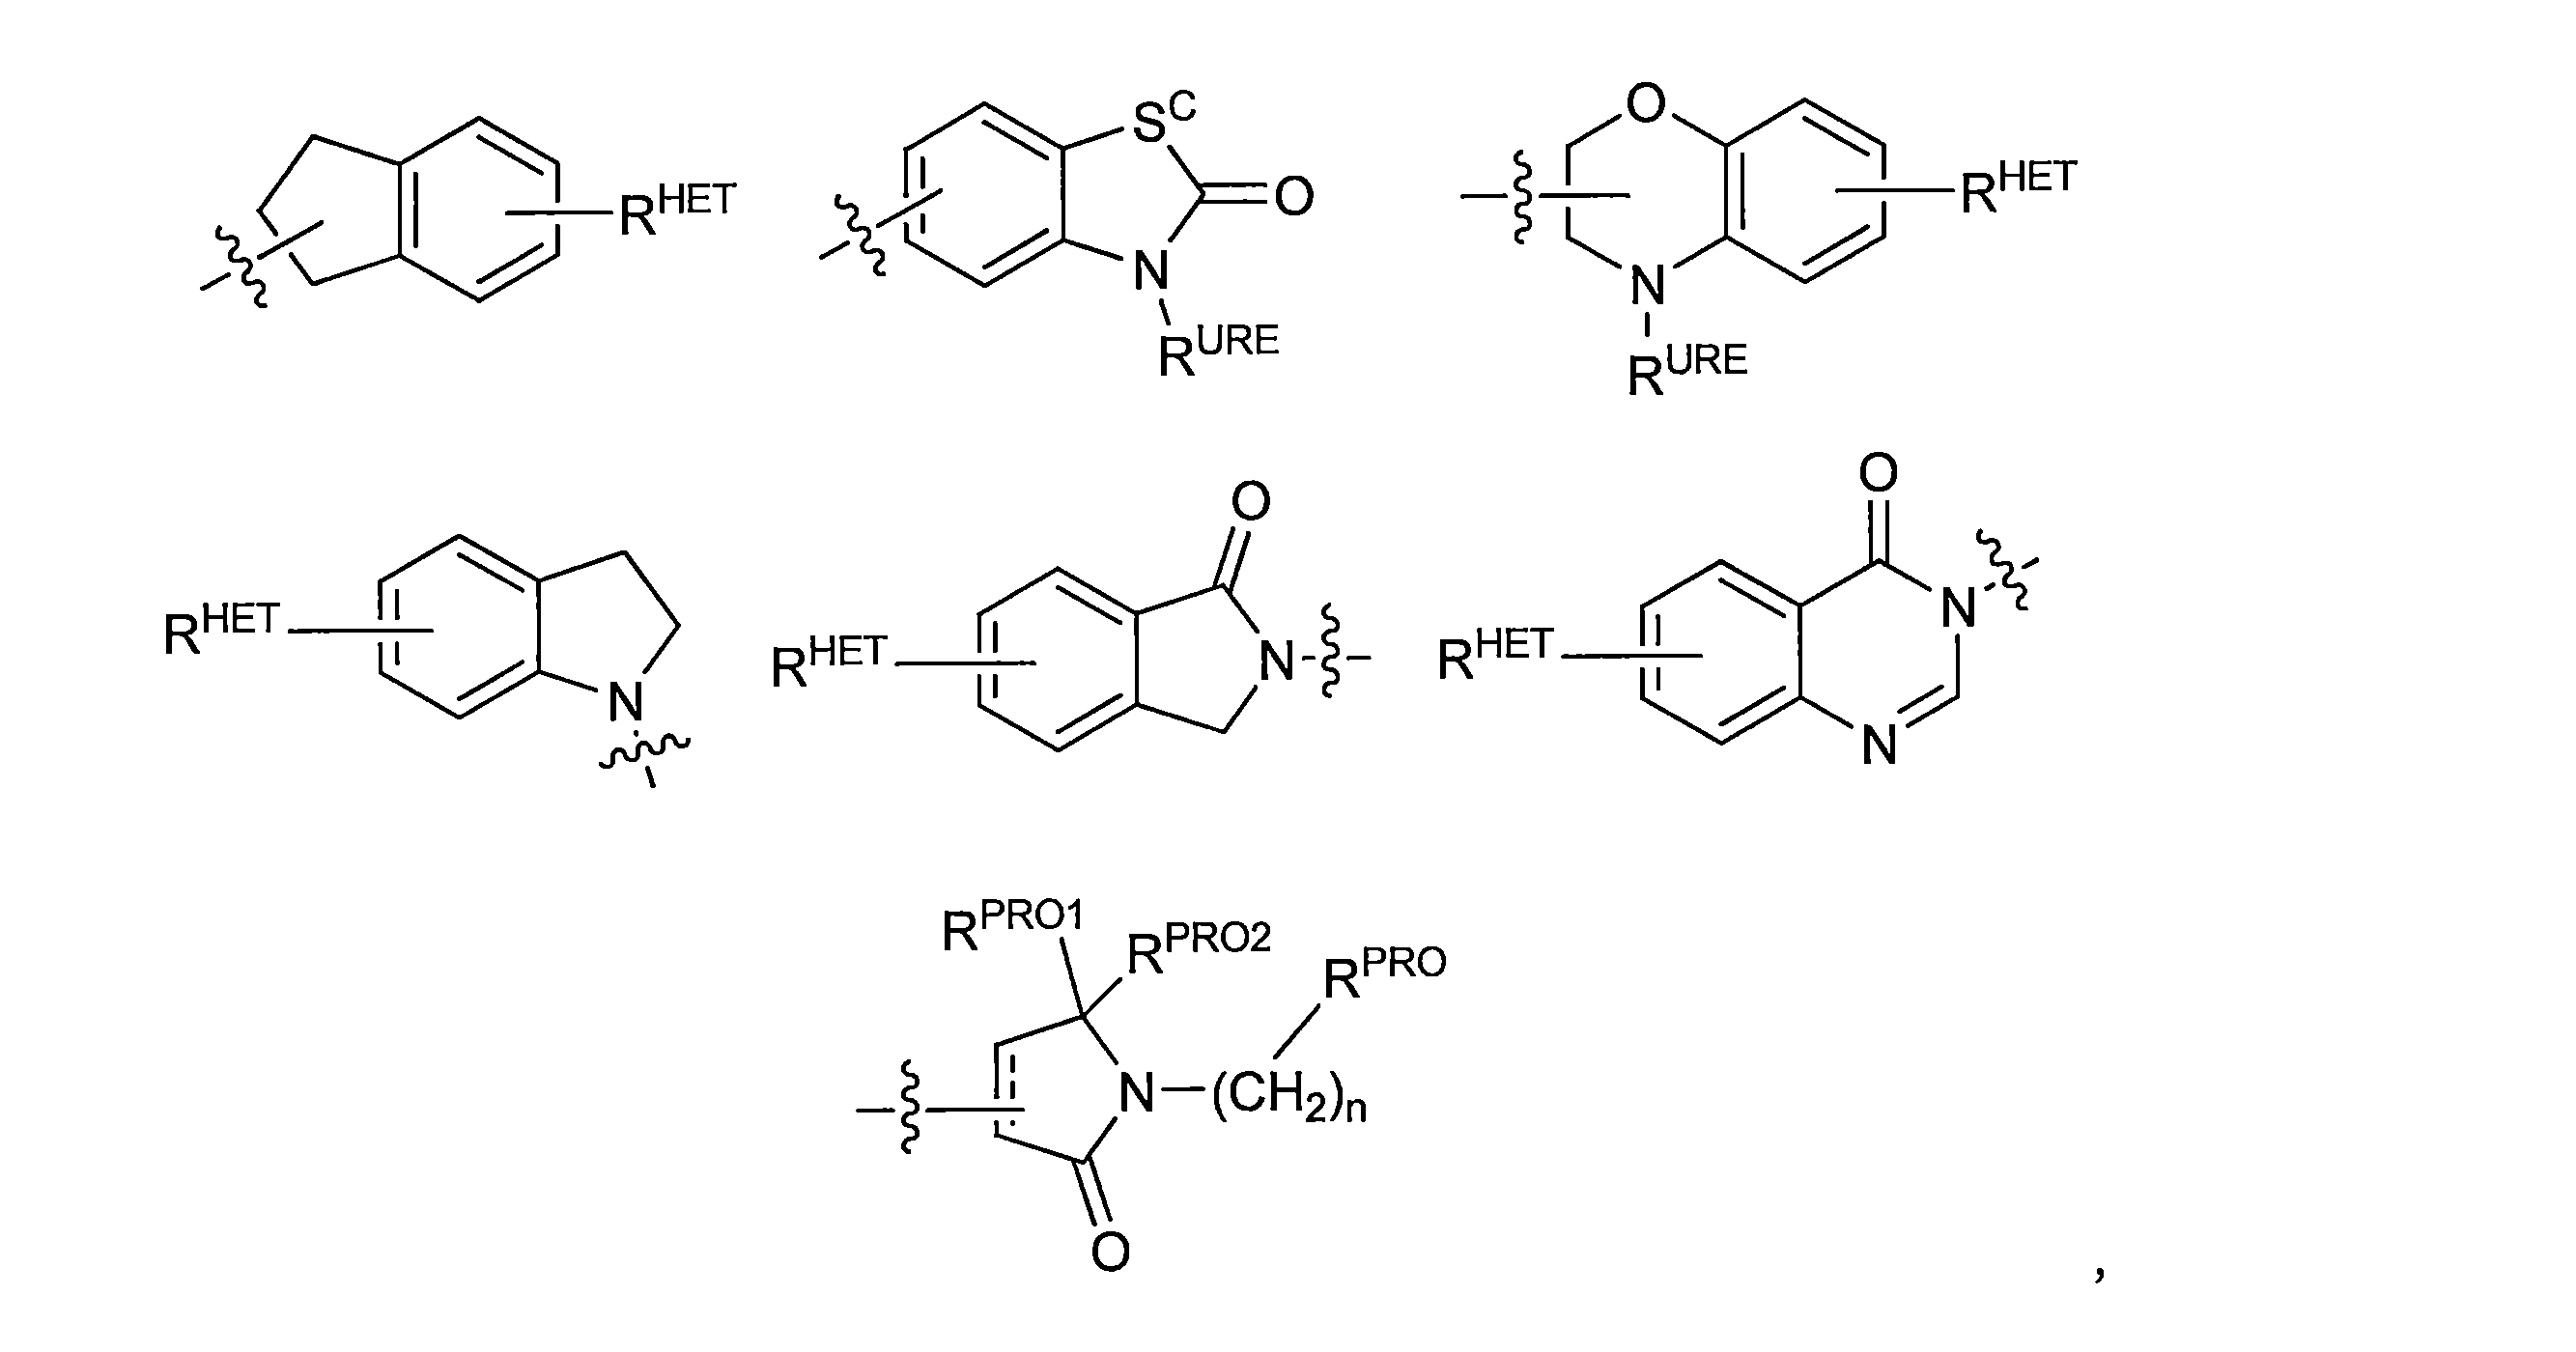
ULM-g~ULM-iのScは、CHRSS、NRUREまたはOであり、
ULM-g~ULM-iのRHETは、H、CN、NO2、ハロ(好ましくはClまたはF)、任意に置換された(好ましくは1個もしくは2個のヒドロキシル基または最大3個のハロ基で置換された)C1-C6アルキル(例えば、CF3)、任意に置換された(好ましくは1個もしくは2個のヒドロキシル基または最大3個のハロ基で置換された)O(C1-C6アルキル)、または任意に置換されたアセチレン基-C≡C-Raであり、式中、Raが、HまたはC1-C6アルキル基(好ましくはC1-C3アルキル)であり、
ULM-g~ULM-iのRSSは、H、CN、NO2、ハロ(好ましくはFまたはCl)、任意に置換されたC1-C6アルキル(好ましくは1つもしくは2つのヒドロキシル基または最大で3個のハロ基で置換される)、任意に置換されたO-(C1-C6アルキル)(好ましくは1つもしくは2つのヒドロキシル基または最大で3個のハロ基で置換される)、または任意に置換された-C(O)(C1-C6アルキル)(好ましくは1つもしくは2つのヒドロキシル基または最大で3個のハロ基で置換される)であり、
ULM-g~ULM-iのRUREは、H、C1-C6アルキル(好ましくはHまたはC1-C3アルキル)、または-C(O)(C1-C6アルキル)であり、それらの基の各々が、1個もしくは2個のヒドロキシル基または最大3個のハロゲン、好ましくはフッ素基で任意に置換されるか、または任意に置換されたフェニル基、任意に置換されたヘテロアリール、または任意に置換されたヘテロシクリル、好ましくは、例えば、ピペリジン、モルホリン、ピロリジン、テトラヒドロフランであり、
ULM-g~ULM-iのRPROは、H、任意に置換されたC1-C6アルキル、またはオキサゾール、イソキサゾール、チアゾール、イソチアゾール、イミダゾール、ジアゾール、オキシイミダゾール、ピロール、ピロリジン、フラン、ジヒドロフラン、テトラヒドロフラン、チエン、ジヒドロチエン、テトラヒドロチエン、ピリジン、ピペリジン、ピペラジン、モルホリン、キノリン、(各々が、好ましくはC1-C3アルキル基で置換され、好ましくはメチルまたはハロ基、好ましくはFまたはCl)、ベンゾフラン、インドール、インドリジン、アザインドリジンからなる群から選択される任意に置換されたアリール(フェニルまたはナプチル)、ヘテロアリール、もしくはヘテロシクリル基であり、
ULM-g~ULM-iのRPRO1およびRPRO2は、各々独立して、H、任意に置換されたC1-C3アルキル基であるか、または一緒になってケト基を形成し、
ULM-g~ULM-iの各nは、独立して、0、1、2、3、4、5、もしくは6(好ましくは0または1)、または任意に置換されたヘテロシクリル、好ましくはテトラヒドロフラン、テトラヒドロチエン、ピペリジン、ピペラジン、もしくはモルホリン(それらの基の各々が、置換された場合、好ましくはメチルまたはハロ(F、Br、Cl)で置換され、それらの基の各々が、リンカー基を介してPTM基(ULM’基を含む)に任意に結合し得る。
Preferred aryl groups for R2' of ULM-g through ULM-i include optionally substituted phenyl or naphthyl groups, preferably phenyl groups, where the phenyl or naphthyl group is linked to a PTM (including a ULM' group) with a linker group, and/or optionally a halogen (preferably F or Cl), an amine, a mono- or di-alkylamine (preferably dimethylamine), F, Cl, OH, COOH, C1-C6 alkyl, preferably CH3, CF3, OMe, OCF3, NO2, or CN group (each of which is a substituted or unsubstituted aryl group in the phenyl ring). and/or a naphthyl group substituted with at least one of a F, Cl, OH, COOH, CH3, CF3, OMe, OCF3, NO2, or CN group (at the ortho, meta, and/or para positions of the phenyl ring, preferably the para position), which may be optionally substituted; an optionally substituted phenyl group (which itself is optionally linked to a PTM group including a ULM' having a linker group); and/or a naphthyl group substituted with at least one of a F, Cl, OH, COOH, CH3, CF3, OMe, OCF3, NO2, or CN group (at the ortho, meta, and/or para positions of the phenyl ring, preferably the para position), which may be optionally substituted; optionally substituted isoxazoles including methyl substituted oxazoles, optionally substituted thiazoles including methyl substituted thiazoles, optionally substituted isothiazoles including methyl substituted isothiazoles, optionally substituted pyrroles including methyl substituted pyrroles, optionally substituted imidazoles including methyl imidazoles, optionally substituted benzimidazoles or methoxybenzylimidazoles, optionally substituted oximidazoles or methyloximidazoles, optionally substituted substituted diazole groups including methyl diazole groups, optionally substituted triazole groups including methyl substituted triazole groups, optionally substituted pyridine groups including halo (preferably F) or methyl substituted pyridine groups, or oxapyridine groups (where the pyridine group is linked to the phenyl group by an oxygen), optionally substituted furans, optionally substituted benzofurans, optionally substituted dihydrobenzofurans, optionally substituted indoles, indolizines, or azaindolizines (2, 3, or 4-azaindolizines), optionally substituted quinolines, optionally substituted groups according to the chemical structure
Sc of ULM-g to ULM-i is CHRSS, NRURE or O;
RHET in ULM-g to ULM-i is H, CN, NO2, halo (preferably Cl or F), optionally substituted C1-C6 alkyl (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups) (e.g. CF3), optionally substituted O(C1-C6 alkyl) (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups) or an optionally substituted acetylenic group -C≡C-Ra, where Ra is H or a C1-C6 alkyl group (preferably C1-C3 alkyl);
RSS of ULM-g through ULM-i is H, CN, NO2, halo (preferably F or Cl), optionally substituted C1-C6 alkyl (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups), optionally substituted O-(C1-C6 alkyl) (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups), or optionally substituted -C(O)(C1-C6 alkyl) (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups);
RURE in ULM-g to ULM-i is H, C1-C6 alkyl (preferably H or C1-C3 alkyl), or -C(O)(C1-C6 alkyl), each of which is optionally substituted with one or two hydroxyl groups or up to three halogens, preferably fluorine groups, or an optionally substituted phenyl group, an optionally substituted heteroaryl, or an optionally substituted heterocyclyl, preferably, for example, piperidine, morpholine, pyrrolidine, tetrahydrofuran;
RPRO in ULM-g to ULM-i is H, an optionally substituted C1-C6 alkyl, or an optionally substituted aryl (phenyl or naphthyl), heteroaryl, or heterocyclyl group selected from the group consisting of oxazole, isoxazole, thiazole, isothiazole, imidazole, diazole, oximidazole, pyrrole, pyrrolidine, furan, dihydrofuran, tetrahydrofuran, thien, dihydrothien, tetrahydrothien, pyridine, piperidine, piperazine, morpholine, quinoline, (each of which is preferably substituted with a C1-C3 alkyl group, preferably methyl or a halo group, preferably F or Cl), benzofuran, indole, indolizine, azaindolizine;
RPRO1 and RPRO2 of ULM-g to ULM-i are each independently H, an optionally substituted C1-C3 alkyl group, or taken together form a keto group;
Each n in ULM-g through ULM-i is independently 0, 1, 2, 3, 4, 5, or 6 (preferably 0 or 1), or optionally substituted heterocyclyl, preferably tetrahydrofuran, tetrahydrothiene, piperidine, piperazine, or morpholine (each of which groups, when substituted, is preferably substituted with methyl or halo (F, Br, Cl), each of which groups may optionally be attached to a PTM group (including a ULM' group) via a linker group.
特定の好ましい態様において、ULM-g~ULM-iの


式中、ULM-g~ULM-iのRPROは、上記と同じである。
In certain preferred embodiments, ULM-g to ULM-i
In the formula, RPRO of ULM-g to ULM-i is the same as above.
ULM-g~ULM-iのR2’のための好ましいヘテロアリール基としては、任意に置換されたキノリン(これはファーマコフォアに結合し得るか、またはキノリン環内の任意の炭素原子上で置換され得る)、任意に置換されたインドール、任意に置換されたインドリジン、任意に置換されたアザインドリジン、任意に置換されたベンゾフラン(任意に置換されたベンゾフランを含む)、任意に置換されたイソキサゾール、任意に置換されたチアゾール、任意に置換されたイソチアゾール、任意に置換されたチオフェン、任意に置換されたピリジン(2-、3、または4-ピリジン)、任意に置換されたイミダゾール、任意に置換されたピロール、任意に置換されたジアゾール、任意に置換されたトリアゾール、テトラゾール、任意に置換されたオキシイミダゾール、または以下の化学構造に従う基が挙げられ、

ULM-g~ULM-iのScは、CHRSS、NRUREまたはOであり、
ULM-g~ULM-iのRHETは、H、CN、NO2、ハロ(好ましくはClまたはF)、任意に置換された(好ましくは1個もしくは2個のヒドロキシル基または最大3個のハロ基で置換された)C1-C6アルキル(例えば、CF3)、任意に置換された(好ましくは1個もしくは2個のヒドロキシル基または最大3個のハロ基で置換された)O(C1-C6アルキル)、または任意に置換されたアセチレン基-C≡C-Raであり、式中、ULM-g~ULM-iのRaが、HまたはC1-C6アルキル基(好ましくはC1-C3アルキル)であり、
ULM-g~ULM-iのRSSは、H、CN、NO2、ハロ(好ましくはFまたはCl)、任意に置換されたC1-C6アルキル(好ましくは1つもしくは2つのヒドロキシル基または最大で3個のハロ基で置換される)、任意に置換されたO-(C1-C6アルキル)(好ましくは1つもしくは2つのヒドロキシル基または最大で3個のハロ基で置換される)、または任意に置換された-C(O)(C1-C6アルキル)(好ましくは1つもしくは2つのヒドロキシル基または最大で3個のハロ基で置換される)であり、
ULM-g~ULM-iのRUREは、H、C1-C6アルキル(好ましくはHまたはC1-C3アルキル)、または-C(O)(C1-C6アルキル)であり、それらの基の各々が、1個もしくは2個のヒドロキシル基または最大3個のハロゲン、好ましくはフッ素基で任意に置換されるか、または任意に置換されたヘテロシクリル、例えば、ピペリジン、モルホリン、ピロリジン、テトラヒドロフラン、テトラヒドロチオフェン、ピペリジン、ピペラジンであり、それらの各々が、任意に置換され、
ULM-g~ULM-iのYCは、NまたはC-RYCであり、式中、RYCが、H、OH、CN、NO2、ハロ(好ましくはClまたはF)、任意に置換された(好ましくは1個もしくは2個のヒドロキシル基または最大3個のハロ基で置換された)C1-C6アルキル(例えば、CF3)、任意に置換された(好ましくは1個もしくは2個のヒドロキシル基または最大3個のハロ基で置換された)O(C1-C6アルキル)、または任意に置換されたアセチレン基-C≡C-Raであり、式中、Raが、HまたはC1-C6アルキル基(好ましくはC1-C3アルキル)であり、それらの基の各々が、リンカー基を介してPTM基(ULM’基を含む)に任意に連結され得る。
Preferred heteroaryl groups for R2' of ULM-g through ULM-i include optionally substituted quinoline (which may be attached to the pharmacophore or substituted on any carbon atom within the quinoline ring), optionally substituted indole, optionally substituted indolizine, optionally substituted azaindolizine, optionally substituted benzofuran (including optionally substituted benzofuran), optionally substituted isoxazole, optionally substituted thiazole, optionally substituted isothiazole, optionally substituted thiophene, optionally substituted pyridine (2-, 3, or 4-pyridine), optionally substituted imidazole, optionally substituted pyrrole, optionally substituted diazole, optionally substituted triazole, tetrazole, optionally substituted oximidazole, or a group according to the chemical structure:
Sc of ULM-g to ULM-i is CHRSS, NRURE or O;
RHET in ULM-g to ULM-i is H, CN, NO2, halo (preferably Cl or F), optionally substituted C1-C6 alkyl (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups) (e.g. CF3), optionally substituted O(C1-C6 alkyl) (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups) or an optionally substituted acetylenic group -C≡C-Ra, where Ra in ULM-g to ULM-i is H or a C1-C6 alkyl group (preferably C1-C3 alkyl);
RSS of ULM-g through ULM-i is H, CN, NO2, halo (preferably F or Cl), optionally substituted C1-C6 alkyl (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups), optionally substituted O-(C1-C6 alkyl) (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups), or optionally substituted -C(O)(C1-C6 alkyl) (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups);
RURE in ULM-g to ULM-i is H, C1-C6 alkyl (preferably H or C1-C3 alkyl), or —C(O)(C1-C6 alkyl), each of which is optionally substituted with one or two hydroxyl groups or up to three halogen, preferably fluorine, groups, or an optionally substituted heterocyclyl, such as piperidine, morpholine, pyrrolidine, tetrahydrofuran, tetrahydrothiophene, piperidine, piperazine, each of which is optionally substituted;
YC in ULM-g through ULM-i is N or C-RYC, where RYC is H, OH, CN, NO2, halo (preferably Cl or F), optionally substituted C1-C6 alkyl (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups) (e.g. CF3), optionally substituted O(C1-C6 alkyl) (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups), or an optionally substituted acetylenic group -C≡C-Ra, where Ra is H or a C1-C6 alkyl group (preferably C1-C3 alkyl), each of which groups may be optionally linked to a PTM group (including a ULM' group) via a linker group.
ULM-g~ULM-iのR2’のための好ましいヘテロシクリルヘテロシクリル基としては、テトラヒドロフラン、テトラヒドロチエン、テトラヒドロキノリン、ピペリジン、ピペラジン、ピロリジン、モルホリン、オキサン、またはチアンが挙げられ、それらの基の各々が、任意に置換され得るか、または以下の化学構造に従う基であり、


式中、
ULM-g~ULM-iのRPROは、H、任意に置換されたC1-C6アルキル、または任意に置換されたアリール、ヘテロアリール、もしくはヘテロシクリル基であり、
ULM-g~ULM-iのRPRO1およびRPRO2はそれぞれ独立して、H、任意に置換されたC1-C3アルキル基であるか、またはともにケト基を形成し、および
ULM-g~ULM-iの各nは独立して、0、1、2、3、4、5、または6(多くの場合、0または1)であり、各基のそれぞれがリンカー基を介してPTM基(ULM’基を含む)に任意で結合され得る。
Preferred heterocyclylheterocyclyl groups for R2′ of ULM-g through ULM-i include tetrahydrofuran, tetrahydrothiene, tetrahydroquinoline, piperidine, piperazine, pyrrolidine, morpholine, oxane, or thiane, each of which may be optionally substituted or is a group according to the chemical structure:
In the formula,
RPRO of ULM-g to ULM-i is H, an optionally substituted C1-C6 alkyl, or an optionally substituted aryl, heteroaryl, or heterocyclyl group;
RPRO1 and RPRO2 of ULM-g through ULM-i are each independently H, an optionally substituted C1-C3 alkyl group, or together form a keto group; and each n of ULM-g through ULM-i is independently 0, 1, 2, 3, 4, 5, or 6 (often 0 or 1), each of which may be optionally attached to a PTM group (including a ULM' group) via a linker group.
ULM-g~ULM-iの好ましいR2’置換基としては、本明細書に開示される特定化合物(本明細書および本明細書に付随する図面に開示される特定化合物を含む)において見出されるR2’置換基も具体的に挙げられる(が、開示される特定化合物に限定されない)。これらR2’置換基の各々は、任意の数のR3’置換基と併せて使用されてもよく、それらも本明細書に開示されている。 Preferred R2' substituents for ULM-g through ULM-i also specifically include (but are not limited to) the R2' substituents found in the specific compounds disclosed herein, including the specific compounds disclosed in this specification and in the drawings accompanying this specification. Each of these R2' substituents may be used in conjunction with any number of R3' substituents, also disclosed herein.
ULM-g~ULM-iのR3’は、好ましくは、任意に置換された-NH-T-アリール、任意に置換された-N(C1-C3アルキル)-T-アリール、任意に置換された-NH-T-ヘテロアリール基、任意に置換された-N(C1-C3アルキル)-T-ヘテロアリール、任意に置換された-NH-T-ヘテロシクリル、または任意に置換された-N(C1-C3アルキル)-T-ヘテロシクリルであり、Tは、任意に置換された-(CH2)n-基であり、メチレン基の各々は、1個もしくは2個の置換基で任意に置換され得、好ましくは、ハロゲン、C1-C3アルカリ基、または本明細書に別途記載されるアミノ酸の側鎖、好ましくは任意に置換され得るメチルから選択され、nは、0~6、多くの場合0、1、2、または3、好ましくは0または1である。あるいは、Tはまた、-(CH2O)n-基、-(OCH2)n-基、-(CH2CH2O)n-基、-(OCH2CH2)n-基であり得、それらの基の各々は任意に置換されている。 R3' in ULM-g to ULM-i is preferably an optionally substituted -NH-T-aryl, an optionally substituted -N(C1-C3 alkyl)-T-aryl, an optionally substituted -NH-T-heteroaryl group, an optionally substituted -N(C1-C3 alkyl)-T-heteroaryl, an optionally substituted -NH-T-heterocyclyl, or an optionally substituted -N(C1-C3 alkyl)-T-heterocyclyl, where T is an optionally substituted -(CH2)n- group, each of the methylene groups may be optionally substituted with one or two substituents, preferably selected from halogen, a C1-C3 alkyl group, or the side chain of an amino acid as otherwise described herein, preferably methyl, which may be optionally substituted, and n is 0 to 6, often 0, 1, 2, or 3, preferably 0 or 1. Alternatively, T can also be a -(CH2O)n- group, a -(OCH2)n- group, a -(CH2CH2O)n- group, or a -(OCH2CH2)n- group, each of which is optionally substituted.
ULM-g~ULM-iのR3’のための好ましいアリール基としては、任意に置換されたフェニルまたはナフチル基、好ましくはフェニル基が挙げられ、フェニルまたはナフチル基は、リンカー基を介してPTM基(ULM’基を含む)に任意に連結され、かつ/あるいはハロゲン(好ましくはFまたはCl)、アミン、モノアルキル-もしくはジアルキルアミン(好ましくはジメチルアミン)、アミド基(好ましくは-(CH2)m-NR1C(O)R2基(式中、m、R1、およびR2が、上記と同じである))、ハロ(多くの場合、FまたはCl)、OH、CH3、CF3、OMe、OCF3、NO2、CN、またはS(O)2RS基(RSが、C1-C6アルキル基、任意に置換されたアリール、ヘテロアリール、もしくはヘテロシクリル基、または-(CH2)m(R″)2基である)で任意に置換され、それらの各々は、フェニル環上のオーソ、メタ、および/もしくはパラ位置(好ましくはパラ位置)で置換され得るか、またはアリール(好ましくはフェニル)、ヘテロアリール、もしくはヘテロシクリルである。好ましくは、該置換フェニル基は、任意に置換されたフェニル基(すなわち、置換フェニル基自体が、好ましくは、F、Cl、OH、SH、COOH、CH3、CF3、OMe、OCF3、NO2、CN、またはPTM基(ULM’基を含む)に結合されているリンカー基のうちの少なくとも1つで置換され、この置換が、フェニル環のオーソ、メタ、および/またはパラ位置、好ましくはパラ位置において起こる)、上記ようなものを含む任意に置換され得るナフチル基、任意に置換されたヘテロアリール(好ましくはメチル置換イソキサゾールを含む任意に置換されたイソキサゾール、メチル置換オキサゾールを含む任意に置換されたオキサゾール、メチル置換チアゾールを含む任意に置換されたチアゾール、メチル置換ピロールを含む任意に置換されたピロール、メチルイミダゾールを含む任意に置換されたイミダゾール、ベンジルイミダゾールもしくはメトキシベンジルイミダゾール、オキシイミダゾールもしくはメチルオキシイミダゾール、メチルジアゾール基を含む任意に置換されたジアゾール基、メチル置換トリアゾール基を含む任意に置換されたトリアゾール基、ハロ-(好ましくは、F)もしくはメチル置換ピリジン基を含むピリジン基、またはオキサピリジン基(ここで、ピリジン基は、酸素によってフェニル基に結合されている)、または任意に置換されたヘテロシクリル(テトラヒドロフラン、テトラヒドロチオフェン、ピロリジン、ピペリジン、モルホリン、ピペラジン、テトラヒドラキノリン、オキサン、またはチアンである。アリール、ヘテロアリール、またはヘテロシクリル基の各々は、リンカー基を介してPTM基(ULM’基を含む)に任意に連結され得る。 Preferred aryl groups for R3' in ULM-g to ULM-i include optionally substituted phenyl or naphthyl groups, preferably phenyl groups, optionally linked to the PTM group (including the ULM' group) via a linker group, and/or halogen (preferably F or Cl), amine, monoalkyl- or dialkylamine (preferably dimethylamine), amide group (preferably -(CH2)m-NRC(O)R2 group, where m, R1, and R2 are the same as above), halo (often F or Cl), OH, CH3, CF3, OMe, OCF3, NO2, CN, or is optionally substituted with an S(O)RS group, where RS is a C1-C6 alkyl group, an optionally substituted aryl, heteroaryl, or heterocyclyl group, or a -(CH2)m(R'') group, each of which may be substituted at the ortho, meta, and/or para positions (preferably the para position) on the phenyl ring, or is an aryl (preferably phenyl), heteroaryl, or heterocyclyl. Preferably, the substituted phenyl group is an optionally substituted phenyl group (i.e., the substituted phenyl group itself is preferably F, Cl, OH, SH, COOH, CH3, CF3, OMe, OCF3, NO2, CN, or P at least one of the linker groups attached to the TM group (including the ULM′ group), the substitution occurring at the ortho, meta, and/or para positions of the phenyl ring, preferably the para position); naphthyl groups which may be optionally substituted including those described above; optionally substituted heteroaryls (preferably optionally substituted isoxazoles including methyl substituted isoxazoles, optionally substituted oxazoles including methyl substituted oxazoles, optionally substituted thiazoles including methyl substituted thiazoles, optionally substituted pyrroles including methyl substituted pyrroles, optionally substituted imidazoles including methylimidazoles, benzyl imidazole or methoxybenzylimidazole, oximidazole or methyloximidazole, optionally substituted diazole groups including methyldiazole groups, optionally substituted triazole groups including methyl-substituted triazole groups, pyridine groups including halo- (preferably F) or methyl-substituted pyridine groups, or oxapyridine groups (wherein the pyridine group is bonded to the phenyl group by an oxygen), or optionally substituted heterocyclyl (tetrahydrofuran, tetrahydrothiophene, pyrrolidine, piperidine, morpholine, piperazine, tetrahydraquinoline, oxane, or thiane. Each of the aryl, heteroaryl, or heterocyclyl groups may be optionally linked to a PTM group (including a ULM' group) via a linker group.
ULM-g~ULM-iのR3’のための好ましいヘテロアリール基としては、任意に置換されたキノリン(これはファーマコフォアに結合し得るか、またはキノリン環内の任意の炭素原子上で置換され得る)、任意に置換されたインドール(ジヒドロインドールを含む)、任意に置換されたインドリジン、任意に置換されたアザインドリジン(2、3、または4-アザインドリジン)、任意に置換されたベンズイミダゾール、ベンゾジアゾール、ベンゾキソフラン、任意に置換されたイミダゾール、任意に置換されたイソキサゾール、任意に置換されたオキサゾール(好ましくはメチル置換)、任意に置換されたジアゾール、任意に置換されたトリアゾール、テトラゾール、任意に置換されたベンゾフラン、任意に置換されたチオフェン、任意に置換されたチアゾール(好ましくはメチルおよび/またはチオール置換)、任意に置換されたイソチアゾール、任意に置換されたトリアゾール(好ましくはメチル基、トリイソプロピルシリル基、任意に置換された-(CH2)m-O-C1-C6アルキル基、または任意に置換された-(CH2)m-C(O)-O-C1-C6アルキル基で置換された1,2,3-トリアゾール)、任意に置換されたピリジン(2-、3、または4-ピリジン)、または以下の化学構造に従う基が挙げられ、

ULM-g~ULM-iのScは、CHRSS、NRUREまたはOであり、
ULM-g~ULM-iのRHETは、H、CN、NO2、ハロ(好ましくはClまたはF)、任意に置換された(好ましくは1個もしくは2個のヒドロキシル基または最大3個のハロ基で置換された)C1-C6アルキル(例えば、CF3)、任意に置換された(好ましくは1個もしくは2個のヒドロキシル基または最大3個のハロ基で置換された)O(C1-C6アルキル)、または任意に置換されたアセチレン基-C≡C-Raであり、式中、Raは、HまたはC1-C6アルキル基(好ましくはC1-C3アルキル)であり、
ULM-g~ULM-iのRSSは、H、CN、NO2、ハロ(好ましくはFまたはCl)、任意に置換されたC1-C6アルキル(好ましくは1つもしくは2つのヒドロキシル基または最大で3個のハロ基で置換される)、任意に置換されたO-(C1-C6アルキル)(好ましくは1つもしくは2つのヒドロキシル基または最大で3個のハロ基で置換される)、または任意に置換された-C(O)(C1-C6アルキル)(好ましくは1つもしくは2つのヒドロキシル基または最大で3個のハロ基で置換される)であり、
ULM-g~ULM-iのRUREは、H、C1-C6アルキル(好ましくはHまたはC1-C3アルキル)、または-C(O)(C1-C6アルキル)であり、それらの基の各々が、1個もしくは2個のヒドロキシル基または最大3個のハロゲン、好ましくはフッ素基で任意に置換されるか、または任意に置換されたヘテロシクリル、例えば、ピペリジン、モルホリン、ピロリジン、テトラヒドロフラン、テトラヒドロチオフェン、ピペリジン、ピペラジンであり、それらの各々が、任意に置換され、
ULM-g~ULM-iのYCは、NまたはC-RYCであり、式中、RYCは、H、OH、CN、NO2、ハロ(好ましくはClまたはF)、任意に置換された(好ましくは1個もしくは2個のヒドロキシル基または最大3個のハロ基で置換された)C1-C6アルキル(例えば、CF3)、任意に置換された(好ましくは1個もしくは2個のヒドロキシル基または最大3個のハロ基で置換された)O(C1-C6アルキル)、または任意に置換されたアセチレン基-C≡C-Raであり、式中、Raは、HまたはC1-C6アルキル基(好ましくはC1-C3アルキル)である。該ヘテロアリール基の各々は、リンカー基を介してPTM基(ULM’基を含む)に任意に連結され得る。
Preferred heteroaryl groups for R3' of ULM-g through ULM-i include optionally substituted quinoline (which may be attached to the pharmacophore or substituted on any carbon atom within the quinoline ring), optionally substituted indole (including dihydroindole), optionally substituted indolizine, optionally substituted azaindolizine (2, 3, or 4-azaindolizine), optionally substituted benzimidazole, benzodiazole, benzoxofuran, optionally substituted imidazole, optionally substituted isoxazole, optionally substituted oxazole (preferably methyl substituted), optionally substituted diazo and optionally substituted aryls, optionally substituted triazoles, tetrazoles, optionally substituted benzofurans, optionally substituted thiophenes, optionally substituted thiazoles (preferably methyl and/or thiol substituted), optionally substituted isothiazoles, optionally substituted triazoles (preferably 1,2,3-triazoles substituted with methyl groups, triisopropylsilyl groups, optionally substituted —(CH2)m-O-C1-C6 alkyl groups, or optionally substituted —(CH2)m-C(O)—O-C1-C6 alkyl groups), optionally substituted pyridines (2-, 3, or 4-pyridine), or a group according to the chemical structure:
Sc of ULM-g to ULM-i is CHRSS, NRURE or O;
RHET in ULM-g to ULM-i is H, CN, NO2, halo (preferably Cl or F), optionally substituted C1-C6 alkyl (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups) (e.g. CF3), optionally substituted O(C1-C6 alkyl) (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups) or an optionally substituted acetylenic group -C≡C-Ra, where Ra is H or a C1-C6 alkyl group (preferably C1-C3 alkyl);
RSS of ULM-g through ULM-i is H, CN, NO2, halo (preferably F or Cl), optionally substituted C1-C6 alkyl (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups), optionally substituted O-(C1-C6 alkyl) (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups), or optionally substituted -C(O)(C1-C6 alkyl) (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups);
RURE in ULM-g to ULM-i is H, C1-C6 alkyl (preferably H or C1-C3 alkyl), or —C(O)(C1-C6 alkyl), each of which is optionally substituted with one or two hydroxyl groups or up to three halogen, preferably fluorine, groups, or an optionally substituted heterocyclyl, such as piperidine, morpholine, pyrrolidine, tetrahydrofuran, tetrahydrothiophene, piperidine, piperazine, each of which is optionally substituted;
YC in ULM-g through ULM-i is N or C-RYC, where RYC is H, OH, CN, NO2, halo (preferably Cl or F), optionally substituted C1-C6 alkyl (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups) (e.g. CF3), optionally substituted O(C1-C6 alkyl) (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups), or an optionally substituted acetylenic group -C≡C-Ra, where Ra is H or a C1-C6 alkyl group (preferably C1-C3 alkyl). Each of the heteroaryl groups may optionally be linked to a PTM group (including a ULM' group) via a linker group.
ULM-g~ULM-iのR3’のための好ましいヘテロシクリル基としては、テトラヒドロキノリン、ピペリジン、ピペラジン、ピロリジン、モルホリン、テトラヒドロフラン、テトラヒドロチエン、オキサン、およびチアンが挙げられ、それらの基の各々は、任意に置換され得るか、または以下の化学構造に従う基であり、


ULM-g~ULM-iのRPROは、H、任意に置換されたC1-C6アルキル、またはオキサゾール、イソキサゾール、チアゾール、イソチアゾール、イミダゾール、ジアゾール、オキシイミダゾール、ピロール、ピロリジン、フラン、ジヒドロフラン、テトラヒドロフラン、チエン、ジヒドロチエン、テトラヒドロチエン、ピリジン、ピペリジン、ピペラジン、モルホリン、キノリン、(各々が、好ましくはC1-C3アルキル基で置換され、好ましくはメチルまたはハロ基、好ましくはFまたはCl)、ベンゾフラン、インドール、インドリジン、アザインドリジンからなる群から選択される任意に置換されたアリール(フェニルまたはナプチル)、ヘテロアリール、もしくはヘテロシクリル基であり、
ULM-g~ULM-iのRPRO1およびRPRO2は、各々独立して、H、任意に置換されたC1-C3アルキル基であるか、またはともにケト基を形成し、
ULM-g~ULM-iの各nは、0、1、2、3、4、5、もしくは6(好ましくは0または1)であり、該ヘテロシクリル基の各々は、リンカー基を介してPTM基(ULM’基を含む)に任意に連結され得る。
Preferred heterocyclyl groups for R3' of ULM-g through ULM-i include tetrahydroquinoline, piperidine, piperazine, pyrrolidine, morpholine, tetrahydrofuran, tetrahydrothiene, oxane, and thiane, each of which may be optionally substituted or is a group according to the chemical structure:
RPRO in ULM-g to ULM-i is H, an optionally substituted C1-C6 alkyl, or an optionally substituted aryl (phenyl or naphthyl), heteroaryl, or heterocyclyl group selected from the group consisting of oxazole, isoxazole, thiazole, isothiazole, imidazole, diazole, oximidazole, pyrrole, pyrrolidine, furan, dihydrofuran, tetrahydrofuran, thien, dihydrothien, tetrahydrothien, pyridine, piperidine, piperazine, morpholine, quinoline, (each of which is preferably substituted with a C1-C3 alkyl group, preferably methyl or a halo group, preferably F or Cl), benzofuran, indole, indolizine, azaindolizine;
RPRO1 and RPRO2 of ULM-g to ULM-i are each independently H, an optionally substituted C1-C3 alkyl group, or together form a keto group;
Each n in ULM-g through ULM-i is 0, 1, 2, 3, 4, 5, or 6 (preferably 0 or 1), and each of the heterocyclyl groups may optionally be linked to a PTM group (including a ULM' group) via a linker group.
ULM-g~ULM-iの好ましいR3’置換基としては、本明細書に開示される特定化合物(本明細書および本明細書に付随する図面に開示される特定化合物を含む)において見出されるR3’置換基も具体的に挙げられる(が、開示される特定化合物に限定されない)。これらR3’置換基の各々は、任意の数のR2’置換基と併せて使用されてもよく、それらも本明細書に開示されている。 Preferred R3' substituents for ULM-g through ULM-i also specifically include (but are not limited to) the R3' substituents found in the specific compounds disclosed herein, including the specific compounds disclosed in this specification and in the drawings accompanying this specification. Each of these R3' substituents may be used in conjunction with any number of R2' substituents, also disclosed herein.
ある特定の代替的な好ましい実施形態では、ULM-g~ULM-iのR2’は、任意に置換された-NR1-XR2’-アルキル基、-NR1-XR2’-アリール基、任意に置換された-NR1-XR2’-HET、任意に置換された-NR1-XR2’-アリール-HET、または任意に置換された-NR1-XR2’-HET-アリールであり、
式中、
ULM-g~ULM-iのR1は、HまたはC1-C3アルキル基(好ましくはH)であり、
ULM-g~ULM-iのXR2’は、任意に置換された-CH2)n-、-CH2)n-CH(Xv)=CH(Xv)-(cisまたはtrans)、-(CH2)n-CH≡CH-、-(CH2CH2O)n-、またはC3-C6シクロアルキル基であり、
ULM-g~ULM-iのXvは、H、ハロ、または1個もしくは2個のヒドロキシル基または最大3個のハロゲン基で任意に置換されたC1-C3アルキル基であり、
ULM-g~ULM-iのアルキルは、任意に置換されたC1-C10アルキル(好ましくはC1-C6アルキル)基(ある特定の好ましい実施形態では、アルキル基は、ハロ基、多くの場合、ClまたはBrで末端キャップされる)であり、
ULM-g~ULM-iのアリールは、任意に置換されたフェニル基またはナフチル基であり(好ましくはフェニル基)、
ULM-g~ULM-iのHETは、任意に置換されたオキサゾール、イソキサゾール、チアゾール、イソチアゾール、イミダゾール、ジアゾール、オキシイミダゾール、ピロール、ピロリジン、フラン、ジヒドロフラン、テトラヒドロフラン、チエン、ジヒドロチエン、テトラヒドロチエン、ピリジン、ピぺリジン、ピペラジン、モルホリン、ベンゾフラン、インドール、インドリジン、アザインドリジン、キノリン(置換される場合、各々好ましくはC1-C3アルキル基、好ましくはメチル、またはハロ基、好ましくはFもしくはClで置換される)、または以下の化学構造に従う基であり、

ULM-g~ULM-iのRHETは、H、CN、NO2、ハロ(好ましくはClまたはF)、任意に置換された(好ましくは1個もしくは2個のヒドロキシル基または最大3個のハロ基で置換された)C1-C6アルキル(例えば、CF3)、任意に置換された(好ましくは1個もしくは2個のヒドロキシル基または最大3個のハロ基で置換された)O(C1-C6アルキル)、または任意に置換されたアセチレン基-C≡C-Raであり、式中、Raは、HまたはC1-C6アルキル基(好ましくはC1-C3アルキル)であり、
ULM-g~ULM-iのRSSは、H、CN、NO2、ハロ(好ましくはFまたはCl)、任意に置換されたC1-C6アルキル(好ましくは1つもしくは2つのヒドロキシル基または最大で3個のハロ基で置換される)、任意に置換されたO-(C1-C6アルキル)(好ましくは1つもしくは2つのヒドロキシル基または最大で3個のハロ基で置換される)、または任意に置換された-C(O)(C1-C6アルキル)(好ましくは1つもしくは2つのヒドロキシル基または最大で3個のハロ基で置換される)であり、
ULM-g~ULM-iのRUREは、H、C1-C6アルキル(好ましくはHまたはC1-C3アルキル)、または-C(O)(C1-C6アルキル)であり、それらの基の各々は、1個もしくは2個のヒドロキシル基または最大3個のハロゲン、好ましくはフッ素基で任意に置換されるか、または任意に置換されたヘテロシクリル、例えば、ピペリジン、モルホリン、ピロリジン、テトラヒドロフラン、テトラヒドロチオフェン、ピペリジン、ピペラジンであり、それらの各々は、任意に置換され、
ULM-g~ULM-iのYCは、NまたはC-RYCであり、式中、RYCは、H、OH、CN、NO2、ハロ(好ましくはClまたはF)、任意に置換された(好ましくは1個もしくは2個のヒドロキシル基または最大3個のハロ基で置換された)C1-C6アルキル(例えば、CF3)、任意に置換された(好ましくは1個もしくは2個のヒドロキシル基または最大3個のハロ基で置換された)O(C1-C6アルキル)、または任意に置換されたアセチレン基-C≡C-Raであり、式中、Raは、HまたはC1-C6アルキル基(好ましくはC1-C3アルキル)であり、
ULM-g~ULM-iのRPROは、H、任意に置換されたC1-C6アルキル、またはオキサゾール、イソキサゾール、チアゾール、イソチアゾール、イミダゾール、ジアゾール、オキシイミダゾール、ピロール、ピロリジン、フラン、ジヒドロフラン、テトラヒドロフラン、チエン、ジヒドロチエン、テトラヒドロチエン、ピリジン、ピペリジン、ピペラジン、モルホリン、キノリン、(各々が、好ましくはC1-C3アルキル基で置換され、好ましくはメチルまたはハロ基、好ましくはFまたはCl)、ベンゾフラン、インドール、インドリジン、アザインドリジンからなる群から選択される任意に置換されたアリール(フェニルまたはナプチル)、ヘテロアリール、もしくはヘテロシクリル基であり、
ULM-g~ULM-iのRPRO1およびRPRO2は、各々独立して、H、任意に置換されたC1-C3アルキル基であるか、またはともにケト基を形成し、
ULM-g~ULM-iの各nは、独立して、0、1、2、3、4、5、または6(好ましくは0または1)である。
In certain alternative preferred embodiments, R2′ of ULM-g through ULM-i is an optionally substituted —NR1-XR2′-alkyl group, an —NR1-XR2′-aryl group, an optionally substituted —NR1-XR2′-HET, an optionally substituted —NR1-XR2′-aryl-HET, or an optionally substituted —NR1-XR2′-HET-aryl;
In the formula,
R1 of ULM-g to ULM-i is H or a C1-C3 alkyl group (preferably H);
XR2' in ULM-g to ULM-i is an optionally substituted -CH2)n-, -CH2)n-CH(Xv)=CH(Xv)-(cis or trans), -(CH2)n-CH≡CH-, -(CH2CH2O)n-, or a C3-C6 cycloalkyl group;
Xv of ULM-g to ULM-i is H, halo, or a C1-C3 alkyl group optionally substituted with 1 or 2 hydroxyl groups or up to 3 halogen groups;
alkyl of ULM-g through ULM-i is an optionally substituted C1-C10 alkyl (preferably C1-C6 alkyl) group (in certain preferred embodiments the alkyl group is end-capped with a halo group, often Cl or Br);
Aryl in ULM-g to ULM-i is an optionally substituted phenyl or naphthyl group (preferably a phenyl group);
HET in ULM-g through ULM-i is an optionally substituted oxazole, isoxazole, thiazole, isothiazole, imidazole, diazole, oximidazole, pyrrole, pyrrolidine, furan, dihydrofuran, tetrahydrofuran, thien, dihydrothien, tetrahydrothien, pyridine, piperidine, piperazine, morpholine, benzofuran, indole, indolizine, azaindolizine, quinoline (each of which, when substituted, is preferably substituted with a C1-C3 alkyl group, preferably methyl, or a halo group, preferably F or Cl), or a group according to the following chemical structure:
RHET in ULM-g to ULM-i is H, CN, NO2, halo (preferably Cl or F), optionally substituted C1-C6 alkyl (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups) (e.g. CF3), optionally substituted O(C1-C6 alkyl) (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups) or an optionally substituted acetylenic group -C≡C-Ra, where Ra is H or a C1-C6 alkyl group (preferably C1-C3 alkyl);
RSS of ULM-g through ULM-i is H, CN, NO2, halo (preferably F or Cl), optionally substituted C1-C6 alkyl (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups), optionally substituted O-(C1-C6 alkyl) (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups), or optionally substituted -C(O)(C1-C6 alkyl) (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups);
RURE in ULM-g to ULM-i is H, C1-C6 alkyl (preferably H or C1-C3 alkyl), or —C(O)(C1-C6 alkyl), each of which is optionally substituted with one or two hydroxyl groups or up to three halogen, preferably fluorine, groups, or an optionally substituted heterocyclyl, such as piperidine, morpholine, pyrrolidine, tetrahydrofuran, tetrahydrothiophene, piperidine, piperazine, each of which is optionally substituted;
YC in ULM-g to ULM-i is N or C-RYC, where RYC is H, OH, CN, NO2, halo (preferably Cl or F), optionally substituted C1-C6 alkyl (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups) (e.g. CF3), optionally substituted O(C1-C6 alkyl) (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups), or an optionally substituted acetylenic group -C≡C-Ra, where Ra is H or a C1-C6 alkyl group (preferably C1-C3 alkyl);
RPRO in ULM-g to ULM-i is H, an optionally substituted C1-C6 alkyl, or an optionally substituted aryl (phenyl or naphthyl), heteroaryl, or heterocyclyl group selected from the group consisting of oxazole, isoxazole, thiazole, isothiazole, imidazole, diazole, oximidazole, pyrrole, pyrrolidine, furan, dihydrofuran, tetrahydrofuran, thien, dihydrothien, tetrahydrothien, pyridine, piperidine, piperazine, morpholine, quinoline, (each of which is preferably substituted with a C1-C3 alkyl group, preferably methyl or a halo group, preferably F or Cl), benzofuran, indole, indolizine, azaindolizine;
RPRO1 and RPRO2 of ULM-g to ULM-i are each independently H, an optionally substituted C1-C3 alkyl group, or together form a keto group;
Each n in ULM-g to ULM-i is independently 0, 1, 2, 3, 4, 5, or 6 (preferably 0 or 1).
該基のそれぞれは、リンカー基を介してPTM基(ULM’基を含む)に任意で結合され得る。 Each of the groups may be optionally attached to a PTM group (including a ULM' group) via a linker group.
本開示のある特定の代替的な好ましい実施形態では、ULM-g~ULM-iのR3’は、任意に置換された-(CH2)n-(V)n’-(CH2)n-(V)n’-RS3’基、任意に置換された-(CH2)n-N(R1’)(C=O)m’-(V)n’-RS3’基、任意に置換された-XR3’-アルキル基、任意に置換された-XR3’-アリール基、任意に置換された-XR3’-HET基、任意に置換された-XR3’-アリール-HET基、または任意に置換された-XR3’-HET-アリール基であり、
式中、
RS3’は、任意に置換されたアルキル基(C1-C10、好ましくはC1-C6アルキル)、任意に置換されたアリール基またはHET基であり、
R1’は、HまたはC1-C3アルキル基(好ましくはH)であり、
VはO、SまたはNR1’であり、
XR3’は、-(CH2)n-、-(CH2CH2O)n-、-CH2)n-CH(Xv)=CH(Xv)-(cisまたはtrans)、-CH2)n-CH≡CH-、またはC3-C6シクロアルキル基であり、すべてが任意に置換され、
Xvは、H、ハロ、または任意で1つもしくは2つのヒドロキシル基または最大で3つのハロゲン基で置換されるC1-C3アルキル基であり、
アルキルは、任意に置換されたC1-C10アルキル(好ましくはC1-C6アルキル)基であり(特定の好ましい実施形態では、アルキル基は、ハロ基、多くの場合はClまたはBrで末端が閉じられている)、
アリールは、任意に置換されたフェニル基またはナフチル基であり(好ましくはフェニル基)、
HETは、任意に置換されたオキサゾール、イソキサゾール、チアゾール、イソチアゾール、イミダゾール、ジアゾール、オキシイミダゾール、ピロール、ピロリジン、フラン、ジヒドロフラン、テトラヒドロフラン、チエン、ジヒドロチエン、テトラヒドロチエン、ピリジン、ピぺリジン、ピペラジン、モルホリン、ベンゾフラン、インドール、インドリジン、アザインドリジン、キノリン(置換される場合、各々好ましくはC1-C3アルキル基、好ましくはメチル、またはハロ基、好ましくはFもしくはClで置換される)、または以下の化学構造に従う基であり、
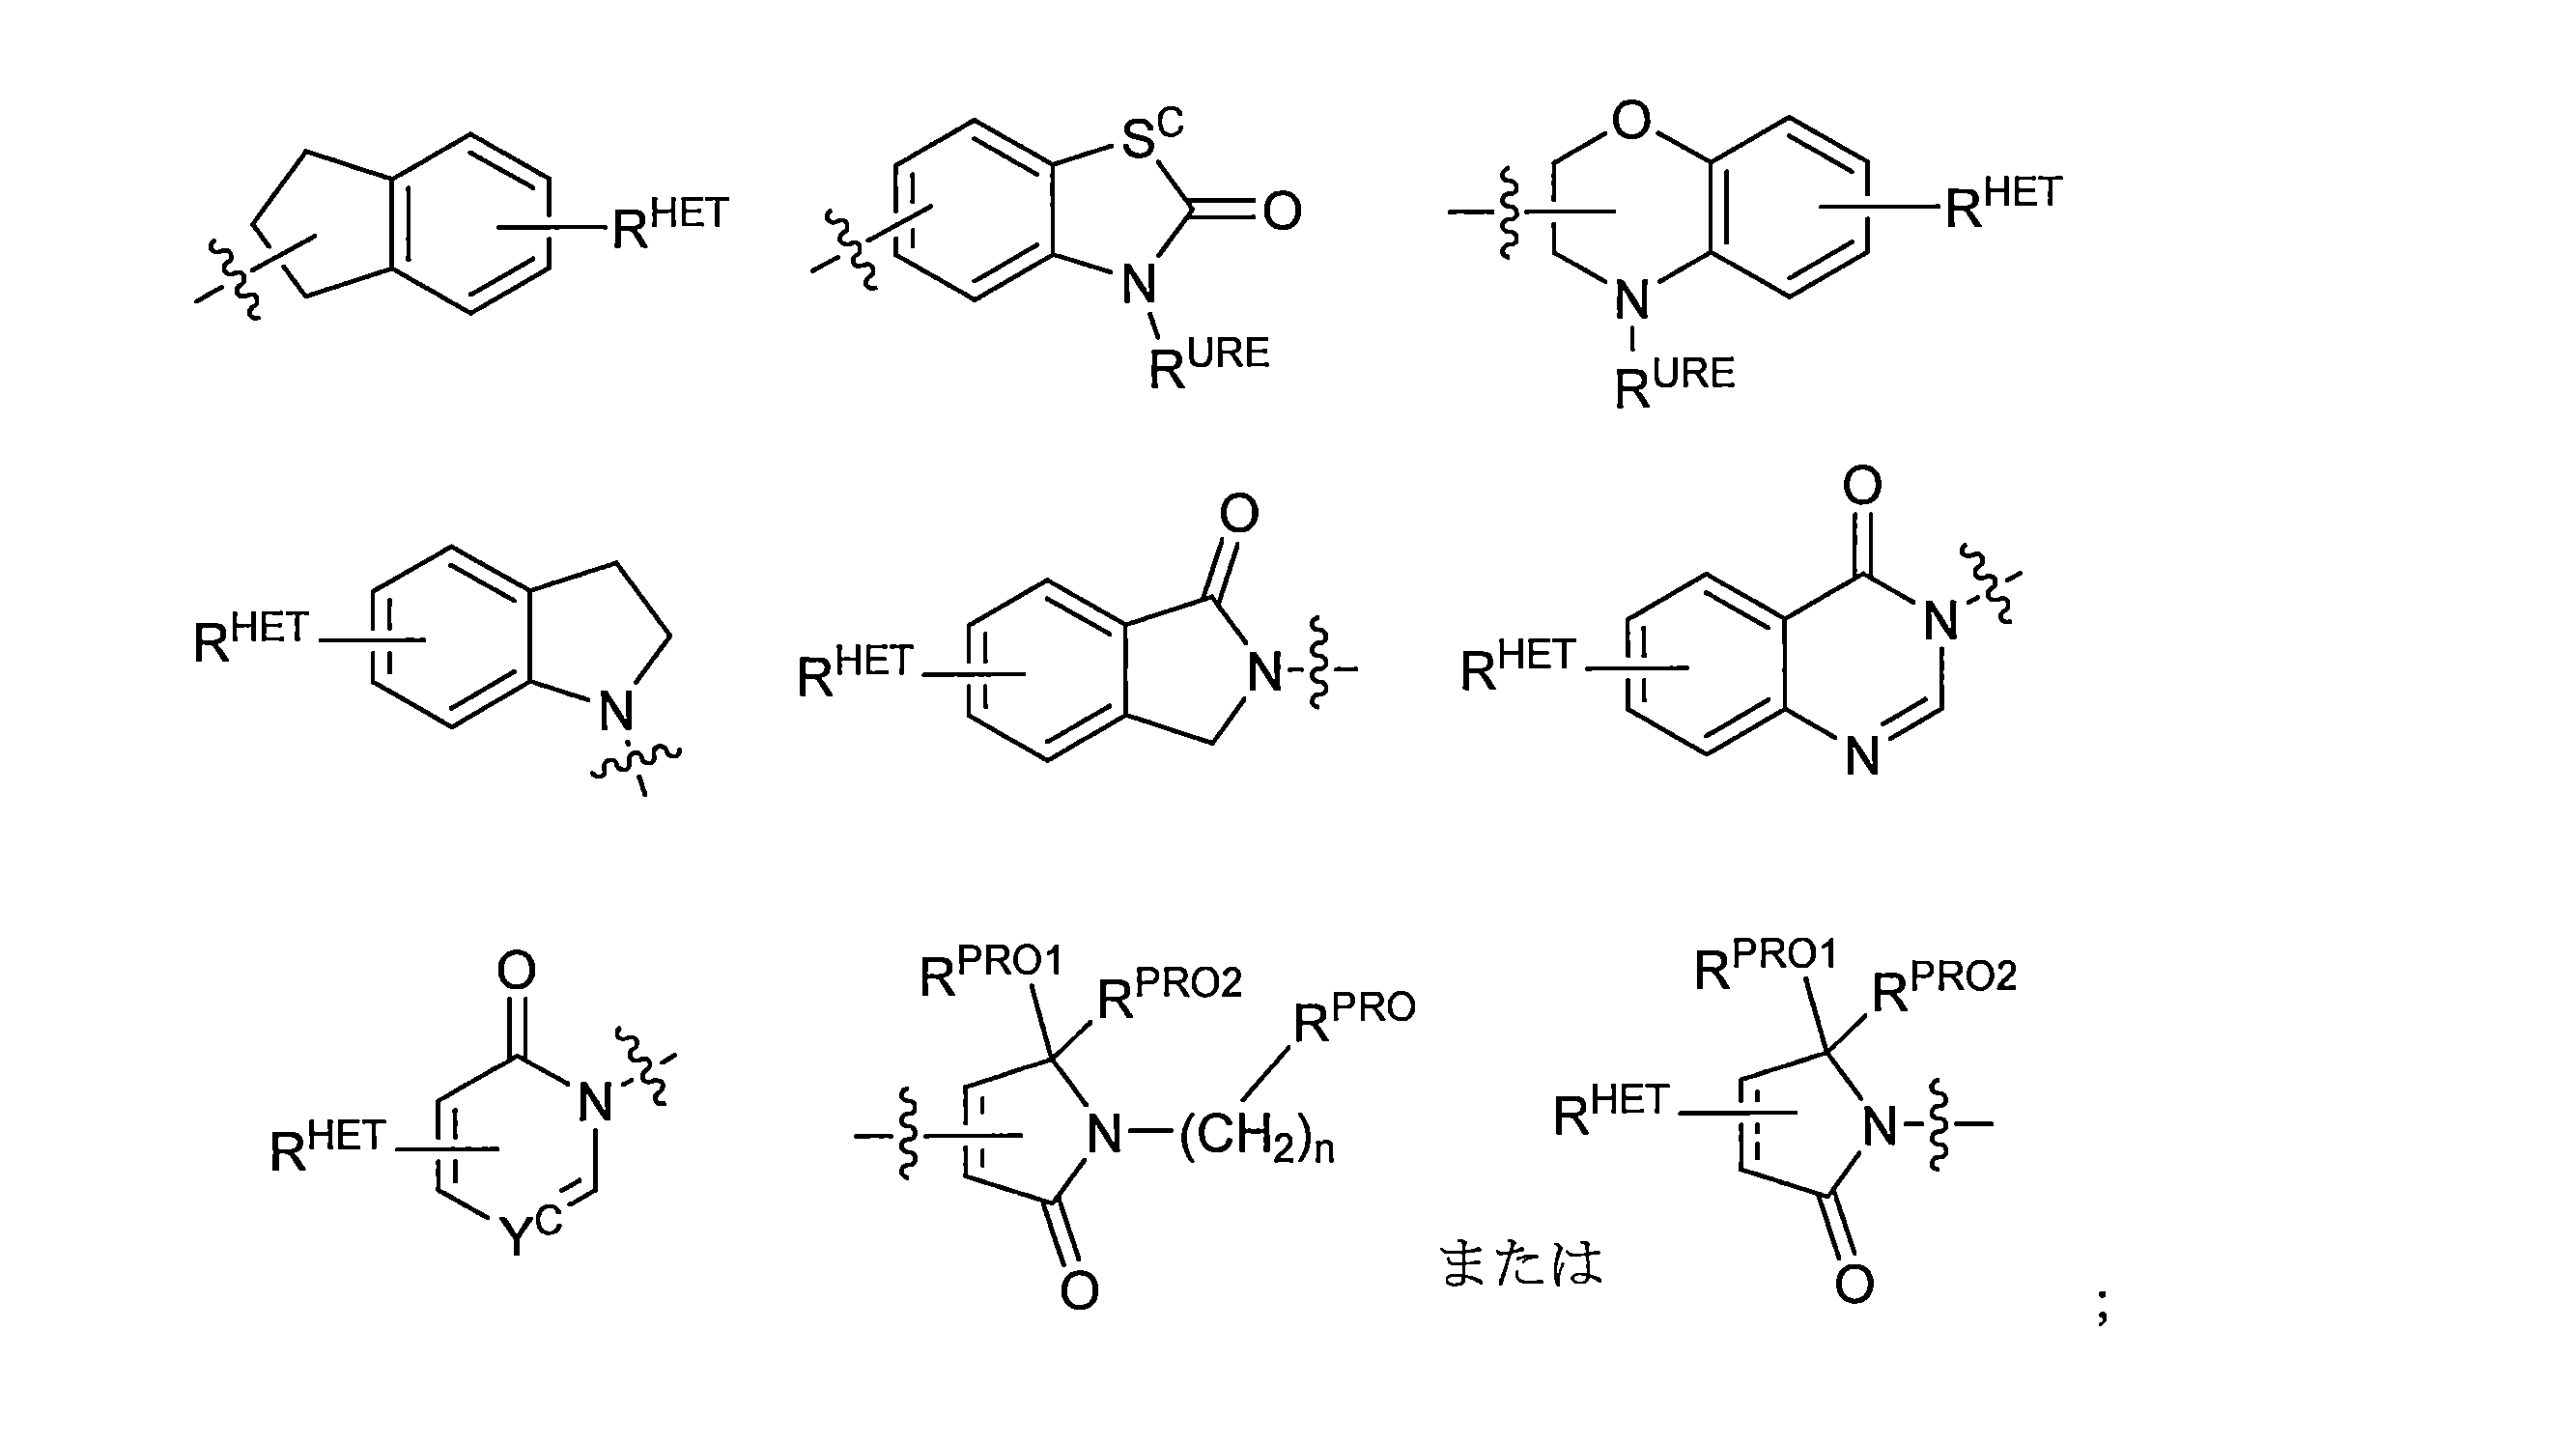
ULM-g~ULM-iのRHETは、H、CN、NO2、ハロ(好ましくはClまたはF)、任意に置換された(好ましくは1個もしくは2個のヒドロキシル基または最大3個のハロ基で置換された)C1-C6アルキル(例えば、CF3)、任意に置換された(好ましくは1個もしくは2個のヒドロキシル基または最大3個のハロ基で置換された)O(C1-C6アルキル)、または任意に置換されたアセチレン基-C≡C-Raであり、式中、Raは、HまたはC1-C6アルキル基(好ましくはC1-C3アルキル)であり、
ULM-g~ULM-iのRSSは、H、CN、NO2、ハロ(好ましくはFまたはCl)、任意に置換されたC1-C6アルキル(好ましくは1つもしくは2つのヒドロキシル基または最大で3個のハロ基で置換される)、任意に置換されたO-(C1-C6アルキル)(好ましくは1つもしくは2つのヒドロキシル基または最大で3個のハロ基で置換される)、または任意に置換された-C(O)(C1-C6アルキル)(好ましくは1つもしくは2つのヒドロキシル基または最大で3個のハロ基で置換される)であり、
ULM-g~ULM-iのRUREは、H、C1-C6アルキル(好ましくはHまたはC1-C3アルキル)、または-C(O)(C0-C6アルキル)であり、それらの基の各々は、1個もしくは2個のヒドロキシル基または最大3個のハロゲン、好ましくはフッ素基で任意に置換されるか、または任意に置換されたヘテロシクリル、例えば、ピペリジン、モルホリン、ピロリジン、テトラヒドロフラン、テトラヒドロチオフェン、ピペリジン、ピペラジンであり、それらの各々は、任意に置換され、
ULM-g~ULM-iのYCは、NまたはC-RYCであり、式中、RYCは、H、OH、CN、NO2、ハロ(好ましくはClまたはF)、任意に置換された(好ましくは1個もしくは2個のヒドロキシル基または最大3個のハロ基で置換された)C1-C6アルキル(例えば、CF3)、任意に置換された(好ましくは1個もしくは2個のヒドロキシル基または最大3個のハロ基で置換された)O(C1-C6アルキル)、または任意に置換されたアセチレン基-C≡C-Raであり、式中、Raは、HまたはC1-C6アルキル基(好ましくはC1-C3アルキル)であり、
ULM-g~ULM-iのRPROは、H、任意に置換されたC1-C6アルキル、またはオキサゾール、イソキサゾール、チアゾール、イソチアゾール、イミダゾール、ジアゾール、オキシイミダゾール、ピロール、ピロリジン、フラン、ジヒドロフラン、テトラヒドロフラン、チエン、ジヒドロチエン、テトラヒドロチエン、ピリジン、ピペリジン、ピペラジン、モルホリン、キノリン、(各々が、好ましくはC1-C3アルキル基で置換され、好ましくはメチルまたはハロ基、好ましくはFまたはCl)、ベンゾフラン、インドール、インドリジン、アザインドリジンからなる群から選択される任意に置換されたアリール(フェニルまたはナプチル)、ヘテロアリール、もしくはヘテロシクリル基であり、
ULM-g~ULM-iのRPRO1およびRPRO2は、各々独立して、H、任意に置換されたC1-C3アルキル基であるか、またはともにケト基を形成し、
ULM-g~ULM-iの各nは、独立して、0、1、2、3、4、5、または6(好ましくは0または1)であり、
ULM-g~ULM-iの各m’は、0または1であり、
ULM-g~ULM-iの各n’は、0または1であり、
式中、該化合物の各々は、好ましくはアルキル基、アリール基またはHet基上で、リンカーを介してPTM基(ULM’基を含む)に任意に連結される。
In certain alternative preferred embodiments of the present disclosure, R3' of ULM-g to ULM-i is an optionally substituted -(CH2)n-(V)n'-(CH2)n-(V)n'-RS3' group, an optionally substituted -(CH2)n-N(R1')(C=O)m'-(V)n'-RS3' group, an optionally substituted -XR3'-alkyl group, an optionally substituted -XR3'-aryl group, an optionally substituted -XR3'-HET group, an optionally substituted -XR3'-aryl-HET group or an optionally substituted -XR3'-HET-aryl group;
In the formula,
RS3' is an optionally substituted alkyl group (C1-C10, preferably C1-C6 alkyl), an optionally substituted aryl group, or a HET group;
R1' is H or a C1-C3 alkyl group (preferably H);
V is O, S or NR1';
XR3' is -(CH2)n-, -(CH2CH2O)n-, -CH2)n-CH(Xv)=CH(Xv)-(cis or trans), -CH2)n-CH≡CH-, or a C3-C6 cycloalkyl group, all optionally substituted;
Xv is H, halo, or a C1-C3 alkyl group optionally substituted with 1 or 2 hydroxyl groups or up to 3 halogen groups;
alkyl is an optionally substituted C1-C10 alkyl (preferably C1-C6 alkyl) group (in certain preferred embodiments, the alkyl group is terminally terminated with a halo group, often Cl or Br);
Aryl is an optionally substituted phenyl or naphthyl group (preferably a phenyl group);
HET is an optionally substituted oxazole, isoxazole, thiazole, isothiazole, imidazole, diazole, oximidazole, pyrrole, pyrrolidine, furan, dihydrofuran, tetrahydrofuran, thien, dihydrothien, tetrahydrothien, pyridine, piperidine, piperazine, morpholine, benzofuran, indole, indolizine, azaindolizine, quinoline (each of which, when substituted, is preferably substituted with a C1-C3 alkyl group, preferably methyl, or a halo group, preferably F or Cl), or a group according to the following chemical structure:
RHET in ULM-g to ULM-i is H, CN, NO2, halo (preferably Cl or F), optionally substituted C1-C6 alkyl (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups) (e.g. CF3), optionally substituted O(C1-C6 alkyl) (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups) or an optionally substituted acetylenic group -C≡C-Ra, where Ra is H or a C1-C6 alkyl group (preferably C1-C3 alkyl);
RSS of ULM-g through ULM-i is H, CN, NO2, halo (preferably F or Cl), optionally substituted C1-C6 alkyl (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups), optionally substituted O-(C1-C6 alkyl) (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups), or optionally substituted -C(O)(C1-C6 alkyl) (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups);
RURE in ULM-g to ULM-i is H, C1-C6 alkyl (preferably H or C1-C3 alkyl), or —C(O)(C0-C6 alkyl), each of which is optionally substituted with one or two hydroxyl groups or up to three halogen, preferably fluorine, groups, or an optionally substituted heterocyclyl, such as piperidine, morpholine, pyrrolidine, tetrahydrofuran, tetrahydrothiophene, piperidine, piperazine, each of which is optionally substituted;
YC in ULM-g to ULM-i is N or C-RYC, where RYC is H, OH, CN, NO2, halo (preferably Cl or F), optionally substituted C1-C6 alkyl (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups) (e.g. CF3), optionally substituted O(C1-C6 alkyl) (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups), or an optionally substituted acetylenic group -C≡C-Ra, where Ra is H or a C1-C6 alkyl group (preferably C1-C3 alkyl);
RPRO in ULM-g to ULM-i is H, an optionally substituted C1-C6 alkyl, or an optionally substituted aryl (phenyl or naphthyl), heteroaryl, or heterocyclyl group selected from the group consisting of oxazole, isoxazole, thiazole, isothiazole, imidazole, diazole, oximidazole, pyrrole, pyrrolidine, furan, dihydrofuran, tetrahydrofuran, thien, dihydrothien, tetrahydrothien, pyridine, piperidine, piperazine, morpholine, quinoline, (each of which is preferably substituted with a C1-C3 alkyl group, preferably methyl or a halo group, preferably F or Cl), benzofuran, indole, indolizine, azaindolizine;
RPRO1 and RPRO2 of ULM-g to ULM-i are each independently H, an optionally substituted C1-C3 alkyl group, or together form a keto group;
Each n in ULM-g to ULM-i is independently 0, 1, 2, 3, 4, 5, or 6 (preferably 0 or 1);
Each m' of ULM-g to ULM-i is 0 or 1,
Each n' of ULM-g to ULM-i is 0 or 1,
wherein each of the compounds is optionally linked, preferably on an alkyl, aryl or Het group, to a PTM group (including a ULM' group) via a linker.
代替的な実施形態では、ULM-g~ULM-iのR3’は、-(CH2)n-アリール、-(CH2CH2O)n-アリール、-(CH2)n-HET、または-(CH2CH2O)n-HETであり、
式中、
ULM-g~ULM-iの該アリールは、1つまたは2つの置換基で任意に置換されたフェニルであり、この場合において当該置換基は、好ましくは-(CH2)nOH、それ自身がCN、ハロ(最大で3つのハロ基)、OH、-(CH2)nO(C1-C6)アルキル、アミン、モノ-またはジ-(C1-C6アルキル)アミンでさらに任意に置換されたC1-C6アルキルから選択され、この場合においてアミンのアルキル基は、1つもしくは2つのヒドロキシル基または最大で3つのハロ(好ましくはF、Cl)基で任意で置換され、または
ULM-g~ULM-iの該アリール基は、-(CH2)nOH、-(CH2)n-O-(C1-C6)アルキル、-(CH2)n-O-(CH2)n-(C1-C6)アルキル、-(CH2)n-C(O)(C0-C6)アルキル、-(CH2)n-C(O)O(C0-C6)アルキル、-(CH2)n-OC(O)(C0-C6)アルキル、アミン、モノ-またはジ-(C1-C6アルキル)アミンで置換され、アミン上のアルキル基は、1個もしくは2個のヒドロキシル基または最大3個のハロ(好ましくはF、Cl)基で任意に置換され、CN、NO2、任意に置換された-(CH2)n-(V)m’-CH2)n-(V)m’-(C1-C6)アルキル基、-(V)m’-(CH2CH2O)n-RPEG基であって、式中、Vは、O、S、またはNR1’であり、R1’は、HまたはC1-C3アルキル基(好ましくはH)であり、RPEGは、H、または任意に置換されたC1-C6アルキル基(カルボキシル基で任意に置換されることを含む)であるか、あるいは
ULM-g~ULM-iの該アリール基は、オキサゾール、イソキサゾール、チアゾール、イソチアゾール、イミダゾール、ジアゾール、オキシイミダゾール、ピロール、ピロリジン、フラン、ジヒドロフラン、テトラヒドロフラン、チエン、ジヒドロチエン、テトラヒドロチエン、ピリジン、ピペリジン、ピペラジン、モルホリン、キノリン、ベンゾフラン、インドール、インドリジン、アザインドリジンからなる群から選択される、ヘテロアリールを含むヘテロシクリルで任意に置換されているか、(置換されている場合、各々が好ましくはC1-C3アルキル基、好ましくはメチルまたはハロ基、好ましくはFまたはClで置換されている)、または以下の化学構造に従う基であり、

ULM-g~ULM-iのRHETは、H、CN、NO2、ハロ(好ましくはClまたはF)、任意に置換された(好ましくは1個もしくは2個のヒドロキシル基または最大3個のハロ基で置換された)C1-C6アルキル(例えば、CF3)、任意に置換された(好ましくは1個もしくは2個のヒドロキシル基または最大3個のハロ基で置換された)O(C1-C6アルキル)、または任意に置換されたアセチレン基-C≡C-Raであり、式中、Raが、HまたはC1-C6アルキル基(好ましくはC1-C3アルキル)であり、
ULM-g~ULM-iのRSSは、H、CN、NO2、ハロ(好ましくはFまたはCl)、任意に置換されたC1-C6アルキル(好ましくは1つもしくは2つのヒドロキシル基または最大で3個のハロ基で置換される)、任意に置換されたO-(C1-C6アルキル)(好ましくは1つもしくは2つのヒドロキシル基または最大で3個のハロ基で置換される)、または任意に置換された-C(O)(C1-C6アルキル)(好ましくは1つもしくは2つのヒドロキシル基または最大で3個のハロ基で置換される)であり、
ULM-g~ULM-iのRUREは、H、C1-C6アルキル(好ましくはHまたはC1-C3アルキル)、または-C(O)(C0-C6アルキル)であり、それらの基の各々が、1個もしくは2個のヒドロキシル基または最大3個のハロゲン、好ましくはフッ素基で任意に置換されるか、または任意に置換されたヘテロシクリル、例えば、ピペリジン、モルホリン、ピロリジン、テトラヒドロフラン、テトラヒドロチオフェン、ピペリジン、ピペラジンであり、それらの各々が、任意に置換され、
ULM-g~ULM-iのYCは、NまたはC-RYCであり、式中、RYCが、H、OH、CN、NO2、ハロ(好ましくはClまたはF)、任意に置換された(好ましくは1個もしくは2個のヒドロキシル基または最大3個のハロ基で置換された)C1-C6アルキル(例えば、CF3)、任意に置換された(好ましくは1個もしくは2個のヒドロキシル基または最大3個のハロ基で置換された)O(C1-C6アルキル)、または任意に置換されたアセチレン基-C≡C-Raであり、式中、Raが、HまたはC1-C6アルキル基(好ましくはC1-C3アルキル)であり、
ULM-g~ULM-iのRPROは、H、任意に置換されたC1-C6アルキル、またはオキサゾール、イソキサゾール、チアゾール、イソチアゾール、イミダゾール、ジアゾール、オキシイミダゾール、ピロール、ピロリジン、フラン、ジヒドロフラン、テトラヒドロフラン、チエン、ジヒドロチエン、テトラヒドロチエン、ピリジン、ピペリジン、ピペラジン、モルホリン、キノリン、(各々が、好ましくはC1-C3アルキル基で置換され、好ましくはメチルまたはハロ基、好ましくはFまたはCl)、ベンゾフラン、インドール、インドリジン、アザインドリジンからなる群から選択される任意に置換されたアリール(フェニルまたはナプチル)、ヘテロアリール、もしくはヘテロシクリル基であり、
ULM-g~ULM-iのRPRO1およびRPRO2はそれぞれ独立して、H、任意に置換されたC1-C3アルキル基であるか、またはともにケト基を形成し、
ULM-g~ULM-iのHETは、好ましくは、オキサゾール、イソキサゾール、チアゾール、イソチアゾール、イミダゾール、ジアゾール、オキシイミダゾール、ピロール、ピロリジン、フラン、ジヒドロフラン、テトラヒドロフラン、チエン、ジヒドロチエン、テトラヒドロチエン、ピリジン、ピぺリジン、ピペラジン、モルホリン、キノリン(各々好ましくはC1-C3アルキル基、好ましくはメチル、またはハロ基、好ましくはFもしくはClで置換される)、ベンゾフラン、インドール、インドリジン、アザインドリジン、または以下の化学構造に従う基であるか、

ULM-g~ULM-iのRHETは、H、CN、NO2、ハロ(好ましくはClまたはF)、任意に置換された(好ましくは1個もしくは2個のヒドロキシル基または最大3個のハロ基で置換された)C1-C6アルキル(例えば、CF3)、任意に置換された(好ましくは1個もしくは2個のヒドロキシル基または最大3個のハロ基で置換された)O(C1-C6アルキル)、または任意に置換されたアセチレン基-C≡C-Raであり、式中、Raが、HまたはC1-C6アルキル基(好ましくはC1-C3アルキル)であり、
ULM-g~ULM-iのRSSは、H、CN、NO2、ハロ(好ましくはFまたはCl)、任意に置換されたC1-C6アルキル(好ましくは1つもしくは2つのヒドロキシル基または最大で3個のハロ基で置換される)、任意に置換されたO-(C1-C6アルキル)(好ましくは1つもしくは2つのヒドロキシル基または最大で3個のハロ基で置換される)、または任意に置換された-C(O)(C1-C6アルキル)(好ましくは1つもしくは2つのヒドロキシル基または最大で3個のハロ基で置換される)であり、
ULM-g~ULM-iのRUREは、H、C1-C6アルキル(好ましくはHまたはC1-C3アルキル)、または-C(O)(C0-C6アルキル)であり、それらの基の各々が、1個もしくは2個のヒドロキシル基または最大3個のハロゲン、好ましくはフッ素基で任意に置換されるか、または任意に置換されたヘテロシクリル、例えば、ピペリジン、モルホリン、ピロリジン、テトラヒドロフラン、テトラヒドロチオフェン、ピペリジン、ピペラジンであり、それらの各々が、任意に置換され、
ULM-g~ULM-iのYCは、NまたはC-RYCであり、式中、RYCは、H、OH、CN、NO2、ハロ(好ましくはClまたはF)、任意に置換された(好ましくは1個もしくは2個のヒドロキシル基または最大3個のハロ基で置換された)C1-C6アルキル(例えば、CF3)、任意に置換された(好ましくは1個もしくは2個のヒドロキシル基または最大3個のハロ基で置換された)O(C1-C6アルキル)、または任意に置換されたアセチレン基-C≡C-Raであり、式中、Raは、HまたはC1-C6アルキル基(好ましくはC1-C3アルキル)であり、
ULM-g~ULM-iのRPROは、H、任意に置換されたC1-C6アルキル、または任意に置換されたアリール、ヘテロアリール、もしくはヘテロシクリル基であり、
ULM-g~ULM-iのRPRO1およびRPRO2はそれぞれ独立して、H、任意に置換されたC1-C3アルキル基であるか、またはともにケト基を形成し、
ULM-g~ULM-iの各m’は独立して、0または1であり、および
ULM-g~ULM-iの各nは独立して、0、1、2、3、4、5、または6(好ましくは0または1)であり、
式中、該化合物のそれぞれは、好ましくは当該アリール基またはHET基上で、任意でリンカー基を介してPTM基(ULM’基を含む)に結合される。
In alternative embodiments, R3' of ULM-g through ULM-i is -(CH2)n-aryl, -(CH2CH2O)n-aryl, -(CH2)n-HET, or -(CH2CH2O)n-HET;
In the formula,
said aryl of ULM-g to ULM-i is phenyl optionally substituted with one or two substituents, where the substituents are preferably selected from -(CH2)nOH, C1-C6 alkyl which itself is optionally further substituted with CN, halo (up to three halo groups), OH, -(CH2)nO(C1-C6)alkyl, amine, mono- or di-(C1-C6 alkyl)amine, where the alkyl group of the amine is optionally substituted with one or two hydroxyl groups or up to three halo (preferably F, Cl) groups; or The aryl group of ULM-g through ULM-i is substituted with -(CH2)nOH, -(CH2)n-O-(C1-C6)alkyl, -(CH2)n-O-(CH2)n-(C1-C6)alkyl, -(CH2)n-C(O)(C0-C6)alkyl, -(CH2)n-C(O)O(C0-C6)alkyl, -(CH2)n-OC(O)(C0-C6)alkyl, amine, mono- or di-(C1-C6 alkyl)amine, the alkyl groups on the amine being substituted with one or two hydroxyl groups or up to optionally substituted with three halo (preferably F, Cl) groups, CN, NO2, an optionally substituted -(CH2)n-(V)m'-CH2)n-(V)m'-(C1-C6) alkyl group, a -(V)m'-(CH2CH2O)n-RPEG group, where V is O, S, or NR1', R1' is H or a C1-C3 alkyl group (preferably H), and RPEG is H or an optionally substituted C1-C6 alkyl group (including optionally substituted with a carboxyl group); The aryl group of ULM-g through ULM-i is optionally substituted with heterocyclyl, including heteroaryl, selected from the group consisting of oxazole, isoxazole, thiazole, isothiazole, imidazole, diazole, oximidazole, pyrrole, pyrrolidine, furan, dihydrofuran, tetrahydrofuran, thien, dihydrothien, tetrahydrothien, pyridine, piperidine, piperazine, morpholine, quinoline, benzofuran, indole, indolizine, azaindolizine (each of which, if substituted, is preferably substituted with a C1-C3 alkyl group, preferably methyl or a halo group, preferably F or Cl), or a group according to the following chemical structure:
RHET in ULM-g to ULM-i is H, CN, NO2, halo (preferably Cl or F), optionally substituted C1-C6 alkyl (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups) (e.g. CF3), optionally substituted O(C1-C6 alkyl) (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups) or an optionally substituted acetylenic group -C≡C-Ra, where Ra is H or a C1-C6 alkyl group (preferably C1-C3 alkyl);
RSS of ULM-g through ULM-i is H, CN, NO2, halo (preferably F or Cl), optionally substituted C1-C6 alkyl (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups), optionally substituted O-(C1-C6 alkyl) (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups), or optionally substituted -C(O)(C1-C6 alkyl) (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups);
RURE in ULM-g to ULM-i is H, C1-C6 alkyl (preferably H or C1-C3 alkyl), or —C(O)(C0-C6 alkyl), each of which is optionally substituted with one or two hydroxyl groups or up to three halogen, preferably fluorine, groups, or an optionally substituted heterocyclyl, such as piperidine, morpholine, pyrrolidine, tetrahydrofuran, tetrahydrothiophene, piperidine, piperazine, each of which is optionally substituted;
YC in ULM-g to ULM-i is N or C-RYC, where RYC is H, OH, CN, NO2, halo (preferably Cl or F), optionally substituted C1-C6 alkyl (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups) (e.g. CF3), optionally substituted O(C1-C6 alkyl) (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups), or an optionally substituted acetylenic group -C≡C-Ra, where Ra is H or a C1-C6 alkyl group (preferably C1-C3 alkyl);
RPRO in ULM-g to ULM-i is H, an optionally substituted C1-C6 alkyl, or an optionally substituted aryl (phenyl or naphthyl), heteroaryl, or heterocyclyl group selected from the group consisting of oxazole, isoxazole, thiazole, isothiazole, imidazole, diazole, oximidazole, pyrrole, pyrrolidine, furan, dihydrofuran, tetrahydrofuran, thien, dihydrothien, tetrahydrothien, pyridine, piperidine, piperazine, morpholine, quinoline, (each of which is preferably substituted with a C1-C3 alkyl group, preferably methyl or a halo group, preferably F or Cl), benzofuran, indole, indolizine, azaindolizine;
RPRO1 and RPRO2 of ULM-g to ULM-i are each independently H, an optionally substituted C1-C3 alkyl group, or together form a keto group;
HET in ULM-g to ULM-i is preferably oxazole, isoxazole, thiazole, isothiazole, imidazole, diazole, oximidazole, pyrrole, pyrrolidine, furan, dihydrofuran, tetrahydrofuran, thien, dihydrothien, tetrahydrothien, pyridine, piperidine, piperazine, morpholine, quinoline (each preferably substituted with a C1-C3 alkyl group, preferably methyl, or a halo group, preferably F or Cl), benzofuran, indole, indolizine, azaindolizine, or a group according to the following chemical structure:
RHET in ULM-g to ULM-i is H, CN, NO2, halo (preferably Cl or F), optionally substituted C1-C6 alkyl (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups) (e.g. CF3), optionally substituted O(C1-C6 alkyl) (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups) or an optionally substituted acetylenic group -C≡C-Ra, where Ra is H or a C1-C6 alkyl group (preferably C1-C3 alkyl);
RSS of ULM-g through ULM-i is H, CN, NO2, halo (preferably F or Cl), optionally substituted C1-C6 alkyl (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups), optionally substituted O-(C1-C6 alkyl) (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups), or optionally substituted -C(O)(C1-C6 alkyl) (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups);
RURE in ULM-g to ULM-i is H, C1-C6 alkyl (preferably H or C1-C3 alkyl), or —C(O)(C0-C6 alkyl), each of which is optionally substituted with one or two hydroxyl groups or up to three halogen, preferably fluorine, groups, or an optionally substituted heterocyclyl, such as piperidine, morpholine, pyrrolidine, tetrahydrofuran, tetrahydrothiophene, piperidine, piperazine, each of which is optionally substituted;
YC in ULM-g to ULM-i is N or C-RYC, where RYC is H, OH, CN, NO2, halo (preferably Cl or F), optionally substituted C1-C6 alkyl (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups) (e.g. CF3), optionally substituted O(C1-C6 alkyl) (preferably substituted with 1 or 2 hydroxyl groups or up to 3 halo groups), or an optionally substituted acetylenic group -C≡C-Ra, where Ra is H or a C1-C6 alkyl group (preferably C1-C3 alkyl);
RPRO of ULM-g to ULM-i is H, an optionally substituted C1-C6 alkyl, or an optionally substituted aryl, heteroaryl, or heterocyclyl group;
RPRO1 and RPRO2 of ULM-g to ULM-i are each independently H, an optionally substituted C1-C3 alkyl group, or together form a keto group;
Each m' of ULM-g through ULM-i is independently 0 or 1, and each n of ULM-g through ULM-i is independently 0, 1, 2, 3, 4, 5, or 6 (preferably 0 or 1);
wherein each of the compounds is preferably attached on the aryl or HET group to a PTM group (including a ULM' group) optionally via a linker group.
さらに追加的実施形態では、好ましい化合物としては以下の化学構造に従うもの、またはその薬学的に許容可能な塩、立体異性体、溶媒和物もしくは多形体が挙げられる:
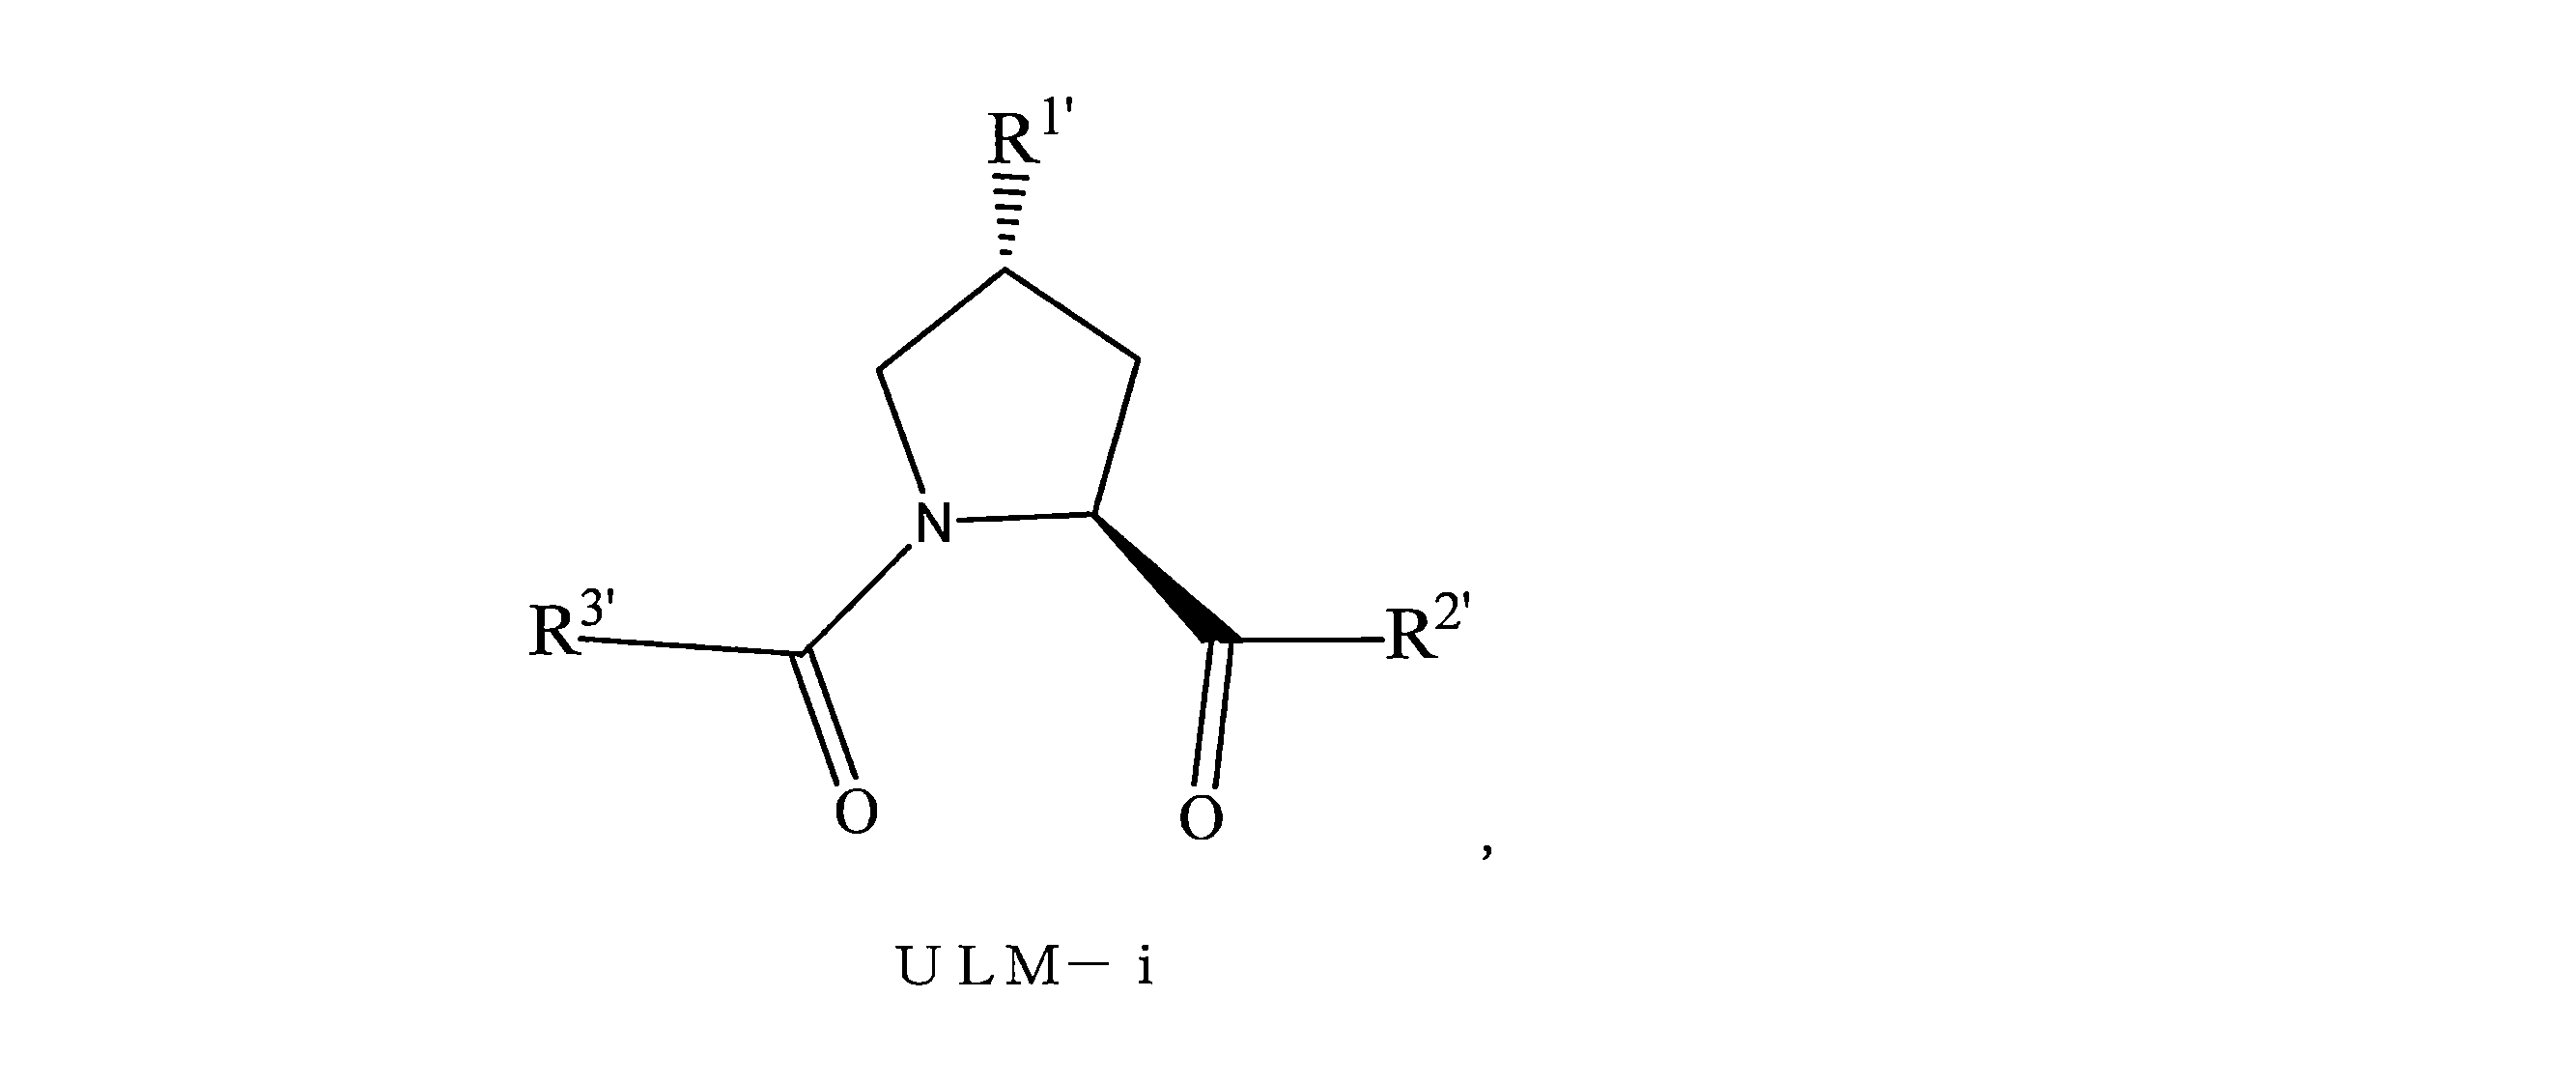
ULM-iのR1’は、OH、または患者もしくは対象においてOHへと代謝される基であり、
ULM-iのR2’は、-NH-CH2-アリール-HET(好ましくは、メチル置換チアゾールに直接結合したフェニル)であり、
ULM-iのR3’は、-CHRCR3’-NH-C(O)-R3P1基または-CHRCR3’-R3P2基であり、
ULM-iのRCR3’は、C1-C4アルキル基、好ましくはメチル、イソプロピル、またはtert-ブチルであり、
ULM-iのR3P1は、C1-C3アルキル(好ましくはメチル)、任意に置換されたオキセタン基(好ましくはメチル置換)、-(CH2)nOCH3基(式中、nが、1または2(好ましくは2)である)、または
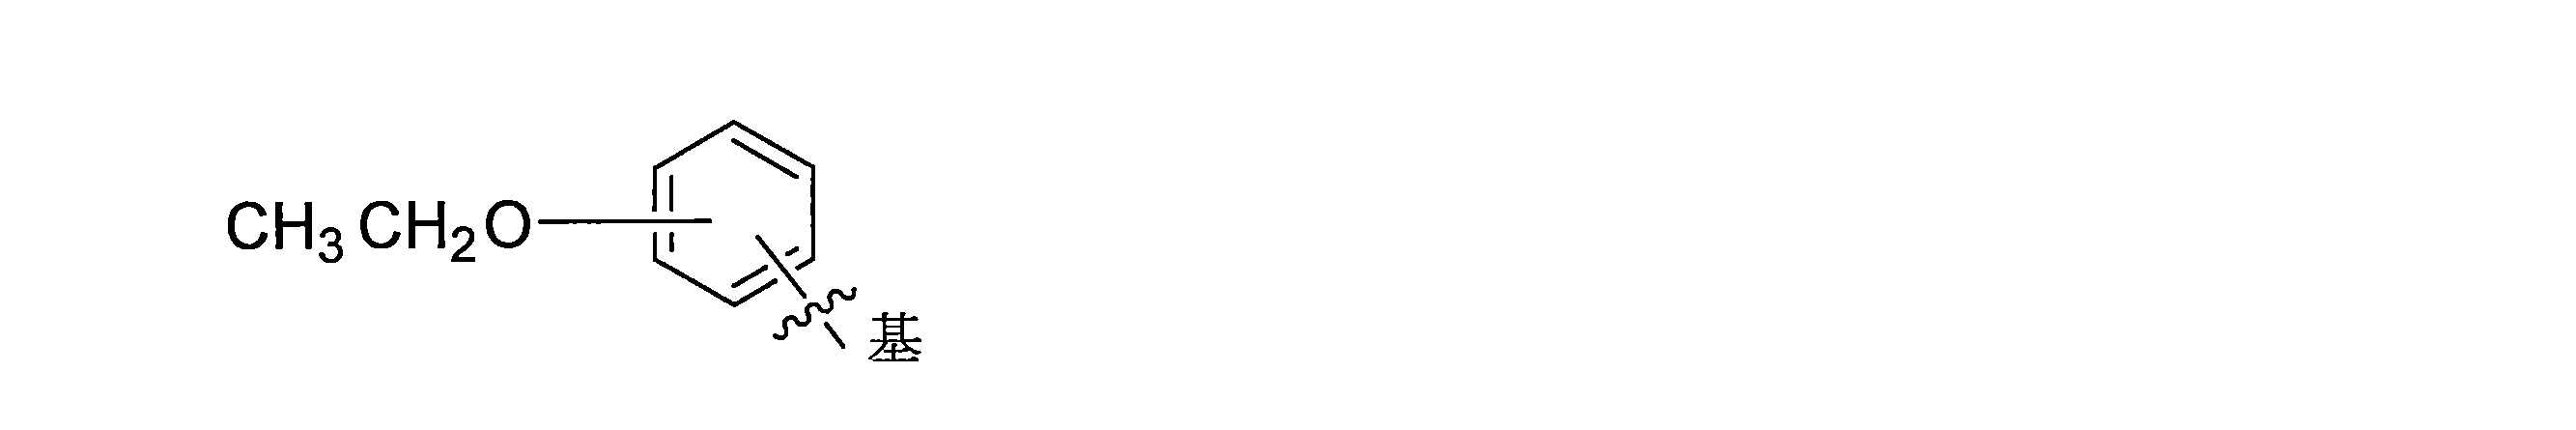
ULM-iのR3P2は、

ULM-iのアリールは、フェニルであり、
ULM-iのHETは、任意に置換されたチアゾールまたはイソチアゾールであり、
ULM-iのRHETは、Hまたはハロ基(好ましくはH)である、化合物、
該化合物は、それぞれリンカー基を介してPTM基(ULM’を含む)に任意で結合される。
In yet additional embodiments, preferred compounds include those according to the following chemical structure, or a pharma- ceutically acceptable salt, stereoisomer, solvate, or polymorph thereof:
R1' of ULM-i is OH or a group that is metabolized to OH in the patient or subject;
R2' of ULM-i is -NH-CH2-aryl-HET (preferably phenyl directly bonded to a methyl substituted thiazole);
R3' of ULM-i is a -CHRCR3'-NH-C(O)-R3P1 group or a -CHRCR3'-R3P2 group;
RCR3' of ULM-i is a C1-C4 alkyl group, preferably methyl, isopropyl, or tert-butyl;
R3P1 of ULM-i is a C1-C3 alkyl (preferably methyl), an optionally substituted oxetane group (preferably methyl substituted), a -(CH2)nOCH3 group (wherein n is 1 or 2 (preferably 2)), or
ULM-i R3P2 is
The aryl of ULM-i is phenyl;
HET of ULM-i is an optionally substituted thiazole or isothiazole;
Compounds in which RHET of ULM-i is H or a halo group (preferably H),
The compounds are optionally attached to the PTM groups (including ULM's) via respective linker groups.
特定の態様において、ユビキチンE3リガーゼ結合部分(ULM)を含む二官能性化合物、またはその薬学的に許容可能な塩、立体異性体、溶媒和物もしくは多形体であって、当該ULMは、以下の化学構造に従う基である:

ULM-jの各R5およびR6は独立して、OH、SH、または任意に置換されたアルキルであるか、またはR5、R6およびそれらが結合する炭素原子でカルボニルを形成し、
ULM-jのR7は、Hまたは任意に置換されたアルキルであり、
ULM-jのEは、結合、C=O、またはC=Sであり、
ULM-jのGは、結合、任意に置換されたアルキル、-COOH、またはC=Jであり、
ULM-jのJは、OまたはN-R8であり、
ULM-jのR8は、H、CN、任意に置換されたアルキル、または任意に置換されたアルコキシであり、
ULM-jのMは、任意に置換されたアリール、任意に置換されたヘテロアリール、任意に置換されたヘテロシクリル、または

ULM-jの各R9およびR10は、独立して、H、任意に置換されたアルキル、任意に置換されたシクロアルキル、任意に置換されたヒドロキシアルキル、任意に置換されたチオアルキル、ULMに結合されたジスルフィド、任意に置換されたヘテロアリール、もしくはハロアルキルであるか、またはR9、R10、およびそれらが結合している炭素原子が、任意に置換されたシクロアルキルを形成し、
ULM-jのR11は、任意に置換されたヘテロシクリル、任意に置換されたアルコキシ、任意に置換されたヘテロアリール、任意に置換されたアリール、または

ULM-jのR12は、Hまたは任意に置換されたアルキルであり、q
ULM-jのR13は、H、任意に置換されたアルキル、任意に置換されたアルキルカルボニル、任意に置換された(シクロアルキル)アルキルカルボニル、任意に置換されたアラルキルカルボニル、任意に置換されたアリールカルボニル、任意に置換された(ヘテロシクリル)カルボニル、または任意に置換されたアラルキル、任意に置換された(オキソアルキル)カルバメートであり、
ULM-jの各R14は、独立して、H、ハロアルキル、任意に置換されたシクロアルキル、任意に置換されたアルキル、アゼチジン、任意に置換されたアルコキシ、または任意に置換されたヘテロシクリルであり、
ULM-jのR15は、H、CN、任意に置換されたヘテロアリール、ハロアルキル、任意に置換されたアリール、任意に置換されたアルコキシ、または任意に置換されたヘテロシクリルであり、
ULM-jの各R16は、独立して、ハロ、任意に置換されたアルキル、任意に置換されたハロアルキル、任意に置換されたCN、または任意に置換されたハロアルコキシであり、
ULM-jの各R25は、独立して、H、もしくは任意に置換されたアルキルであるか、または両方のR25基が一緒になって、オキソもしくは任意に置換されたシクロアルキル基を形成することができ、
ULM-jのR23は、HまたはOHであり、
ULM-jのZ1、Z2、Z3、およびZ4が、独立して、CまたはNであり、
ULM-jのoは、0、1、2、3もしくは4である。
In certain embodiments, a bifunctional compound comprising a ubiquitin E3 ligase binding moiety (ULM), or a pharma- ceutically acceptable salt, stereoisomer, solvate or polymorph thereof, wherein the ULM is a group according to the following chemical structure:
each R5 and R6 of ULM-j is independently OH, SH, or an optionally substituted alkyl, or R5, R6 and the carbon atom to which they are attached form a carbonyl;
R7 of ULM-j is H or optionally substituted alkyl;
E of ULM-j is a bond, C=O, or C=S;
G of ULM-j is a bond, optionally substituted alkyl, —COOH, or C═J;
J of ULM-j is O or N-R8;
R8 of ULM-j is H, CN, optionally substituted alkyl, or optionally substituted alkoxy;
M of ULM-j is optionally substituted aryl, optionally substituted heteroaryl, optionally substituted heterocyclyl, or
each R9 and R10 of ULM-j is independently H, optionally substituted alkyl, optionally substituted cycloalkyl, optionally substituted hydroxyalkyl, optionally substituted thioalkyl, a disulfide bonded to ULM, optionally substituted heteroaryl, or haloalkyl, or R9, R10 and the carbon atom to which they are attached form an optionally substituted cycloalkyl;
R11 of ULM-j is optionally substituted heterocyclyl, optionally substituted alkoxy, optionally substituted heteroaryl, optionally substituted aryl, or
R12 of ULM-j is H or optionally substituted alkyl, and q
R13 of ULM-j is H, optionally substituted alkyl, optionally substituted alkylcarbonyl, optionally substituted (cycloalkyl)alkylcarbonyl, optionally substituted aralkylcarbonyl, optionally substituted arylcarbonyl, optionally substituted (heterocyclyl)carbonyl, or optionally substituted aralkyl, optionally substituted (oxoalkyl)carbamate;
each R14 in ULM-j is independently H, haloalkyl, optionally substituted cycloalkyl, optionally substituted alkyl, azetidine, optionally substituted alkoxy, or optionally substituted heterocyclyl;
R15 of ULM-j is H, CN, optionally substituted heteroaryl, haloalkyl, optionally substituted aryl, optionally substituted alkoxy, or optionally substituted heterocyclyl;
each R16 in ULM-j is independently halo, optionally substituted alkyl, optionally substituted haloalkyl, optionally substituted CN, or optionally substituted haloalkoxy;
Each R25 of ULM-j is independently H, or an optionally substituted alkyl, or both R25 groups together can form an oxo or an optionally substituted cycloalkyl group;
R23 of ULM-j is H or OH;
Z1, Z2, Z3, and Z4 of ULM-j are independently C or N;
o of ULM-j is 0, 1, 2, 3 or 4.
特定の実施形態では、ULM-jのGは、C=Jであり、JはOであり、R7はHであり、各R14はHであり、そしてoは0である。 In certain embodiments, G in ULM-j is C=J, J is O, R7 is H, each R14 is H, and o is 0.
特定の実施形態では、ULM-jのGは、C=Jであり、JはOであり、R7はHであり、各R14はHであり、R15は任意に置換されたヘテロアリールであり、そしてoは0である。他の例では、EはC=Oであり、Mは

ULM-jのEが、C=Oである、ある特定の実施形態では、R11は、任意に置換されたヘテロシクリルまたは

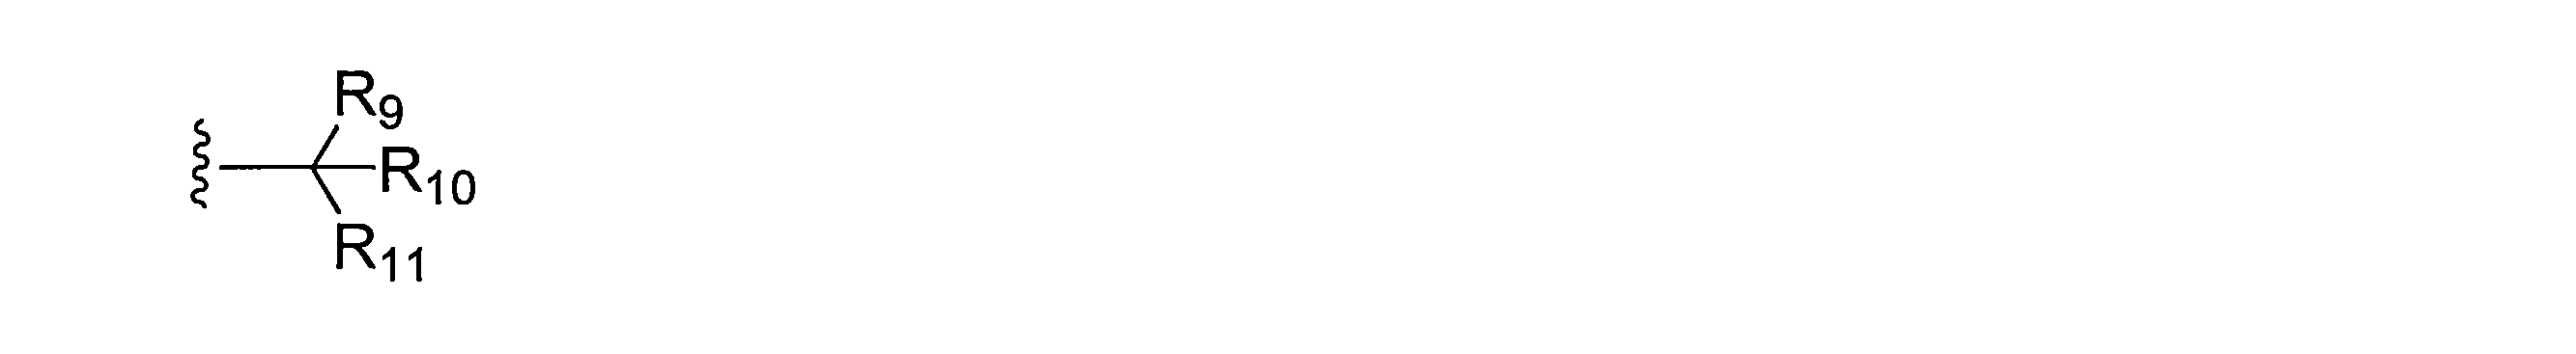
ULM-jのEが、C=Oであり、Mが、

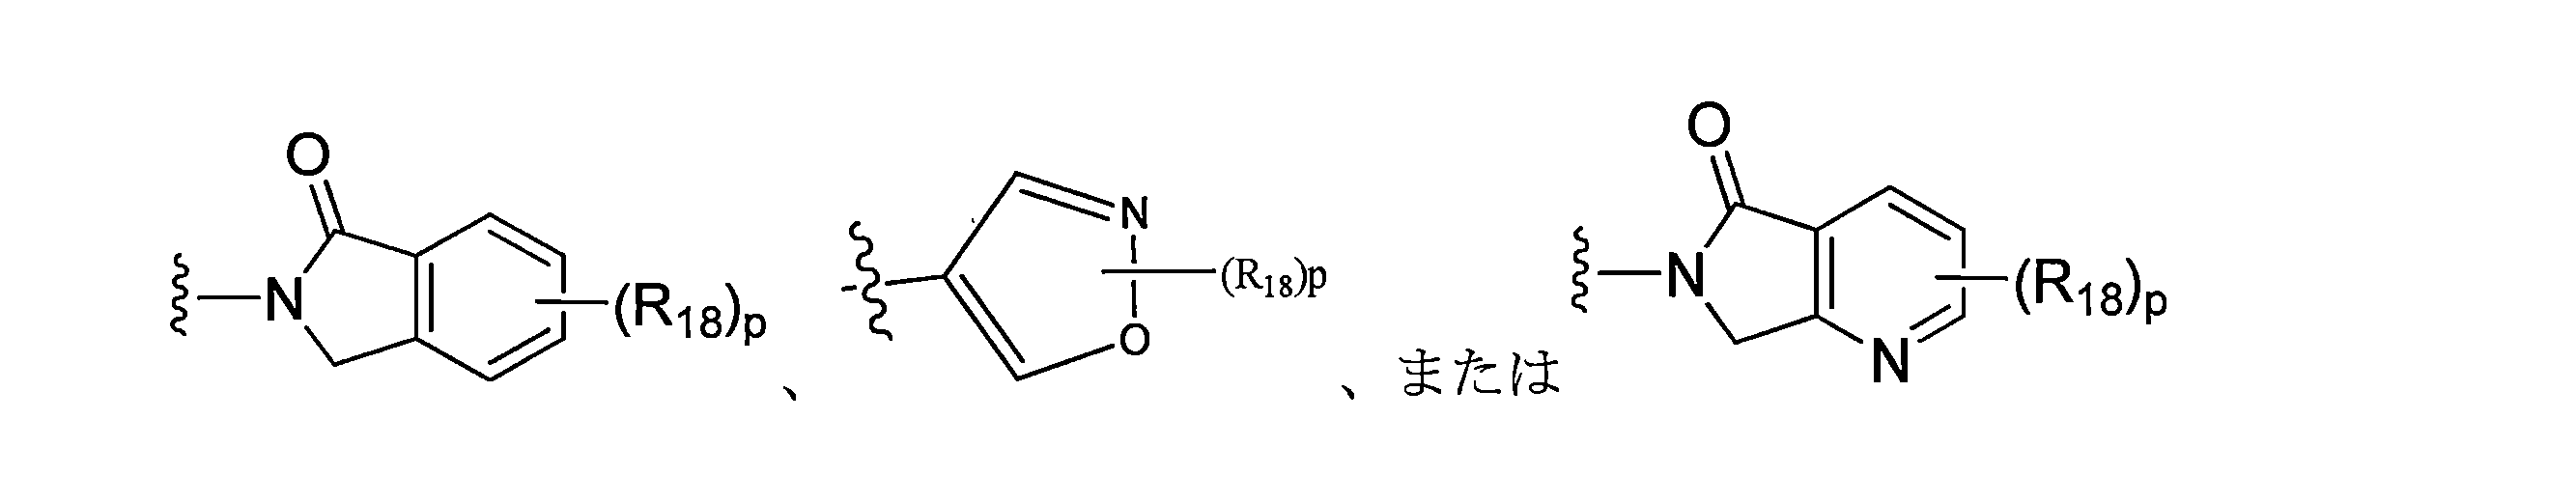
ある特定の実施形態では、各R14は、独立して、H、ヒドロキシル、ハロ、アミン、アミド、アルコキシ、アルキル、ハロアルキル、または複素環のうちの少なくとも1つで置換されている。 In certain embodiments, each R14 is independently substituted with at least one of H, hydroxyl, halo, amine, amide, alkoxy, alkyl, haloalkyl, or heterocycle.
ある特定の実施形態では、ULM-jのR15は、

特定の実施形態では、ULM、および存在する場合にはULM’はそれぞれ独立して、以下の化学構造に従う基である:

ULM-kのGは、C=Jであり、JはOであり、
ULM-kのR7はHであり、
ULM-kの各R14は、独立して、H、アミド、アルキル、例えば、1つ以上のC1-C6アルキル基またはC(O)NR’R”で任意に置換されたメチルであり、
R’およびR”は、各々独立して、H、任意に置換されたアルキル、またはシクロアルキルであり、
ULM-kのoは0であり、
ULM-kのR15は、ULM-jについて上記に定義されるとおりであり、
ULM-kのR16は、ULM-jについて上記に定義されるとおりであり、
ULM-kのR17は、H、ハロ、任意に置換されたシクロアルキル、任意に置換されたアルキル、任意に置換されたアルケニル、およびハロアルキルである。
In certain embodiments, ULM, and, if present, ULM′, are each independently a group conforming to the following chemical structure:
G of ULM-k is C=J, J is O,
R7 of ULM-k is H;
Each R14 in ULM-k is independently H, amido, alkyl, e.g., methyl optionally substituted with one or more C1-C6 alkyl groups or C(O)NR'R";
R′ and R″ are each independently H, optionally substituted alkyl, or cycloalkyl;
o of ULM-k is 0;
R15 of ULM-k is as defined above for ULM-j;
R16 of ULM-k is as defined above for ULM-j;
R17 of ULM-k is H, halo, optionally substituted cycloalkyl, optionally substituted alkyl, optionally substituted alkenyl, and haloalkyl.
他の例では、ULM-kのR17はアルキル(例えば、メチル)またはシクロアルキル(例えば、シクロプロピル)である。 In other examples, R17 in ULM-k is alkyl (e.g., methyl) or cycloalkyl (e.g., cyclopropyl).
他の実施形態では、ULM、および存在する場合にはULM’はそれぞれ独立して、以下の化学構造に従う基である:

ULM-kのGは、C=Jであり、JはOであり、
ULM-kのR7はHであり、
ULM-kの各R14はHであり、
ULM-kのoは0であり、および
ULM-kのR15が、任意に置換された以下からなる群から選択される。
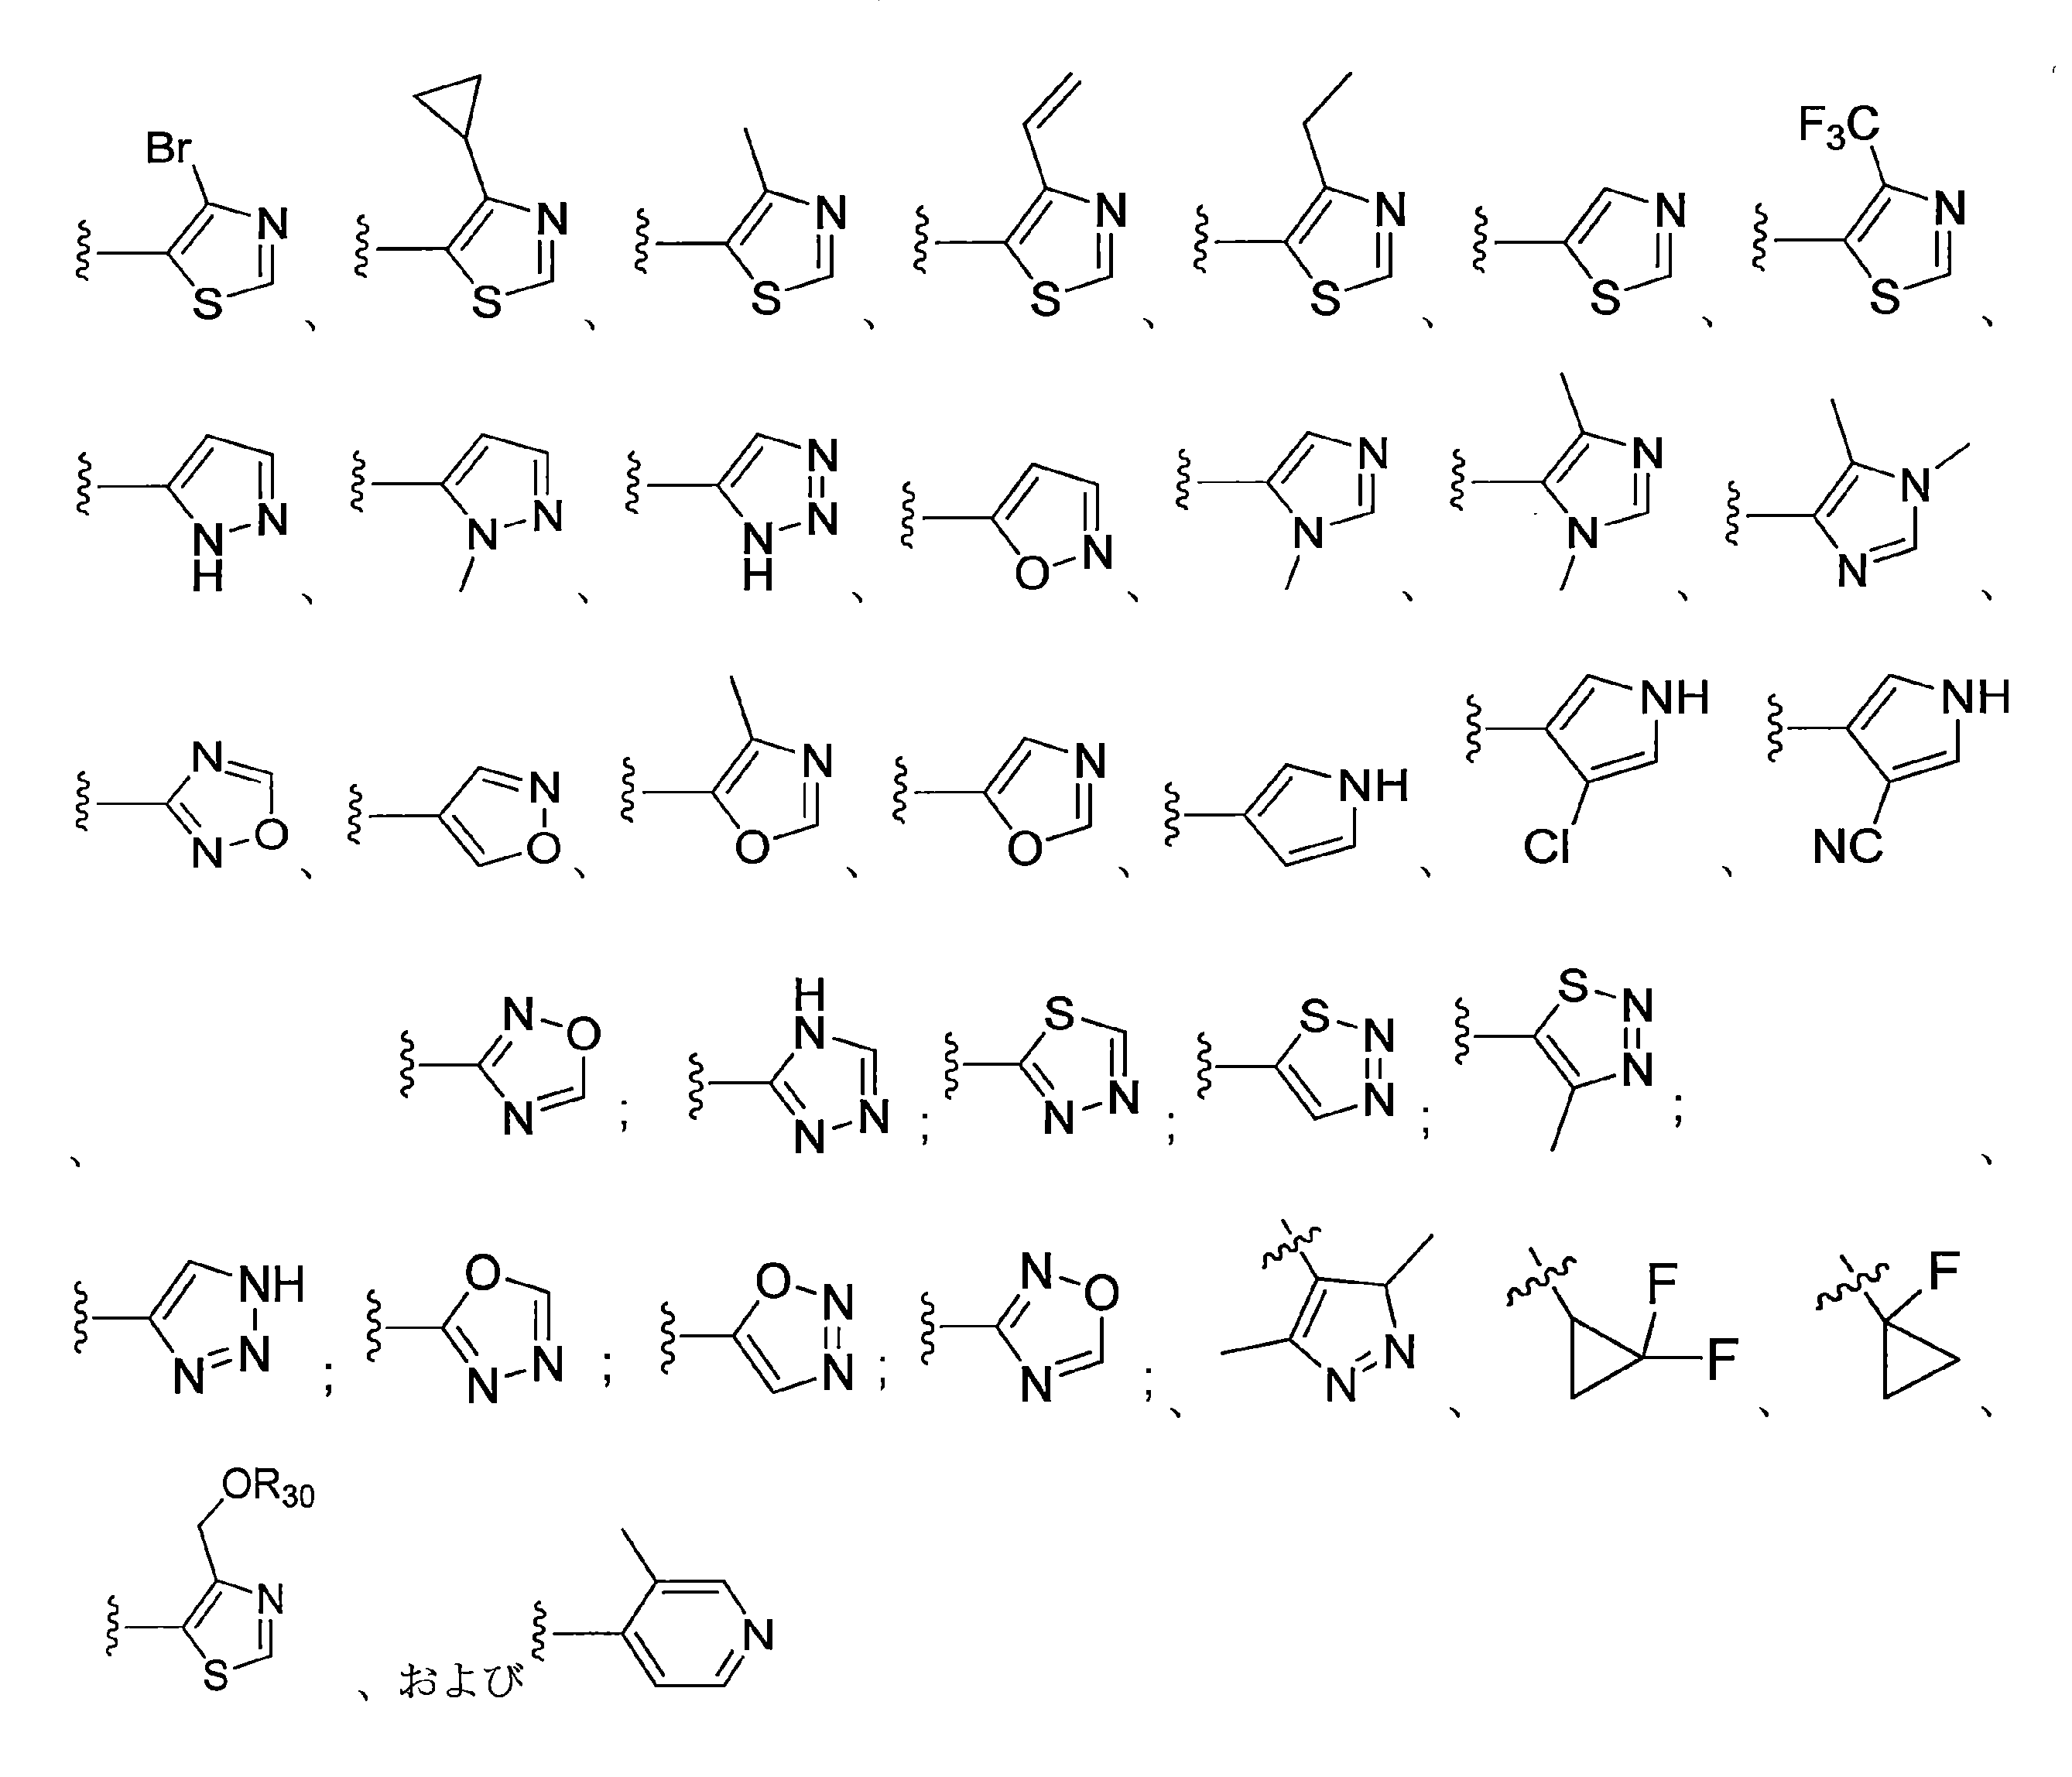
G of ULM-k is C=J, J is O,
R7 of ULM-k is H;
Each R14 in ULM-k is H;
o in ULM-k is 0, and R15 in ULM-k is selected from the group consisting of optionally substituted:
他の実施形態では、ULM、および存在する場合にはULM’はそれぞれ独立して、以下の化学構造に従う基である:

ULM-kのEは、C=Oであり、
ULM-kのMは、
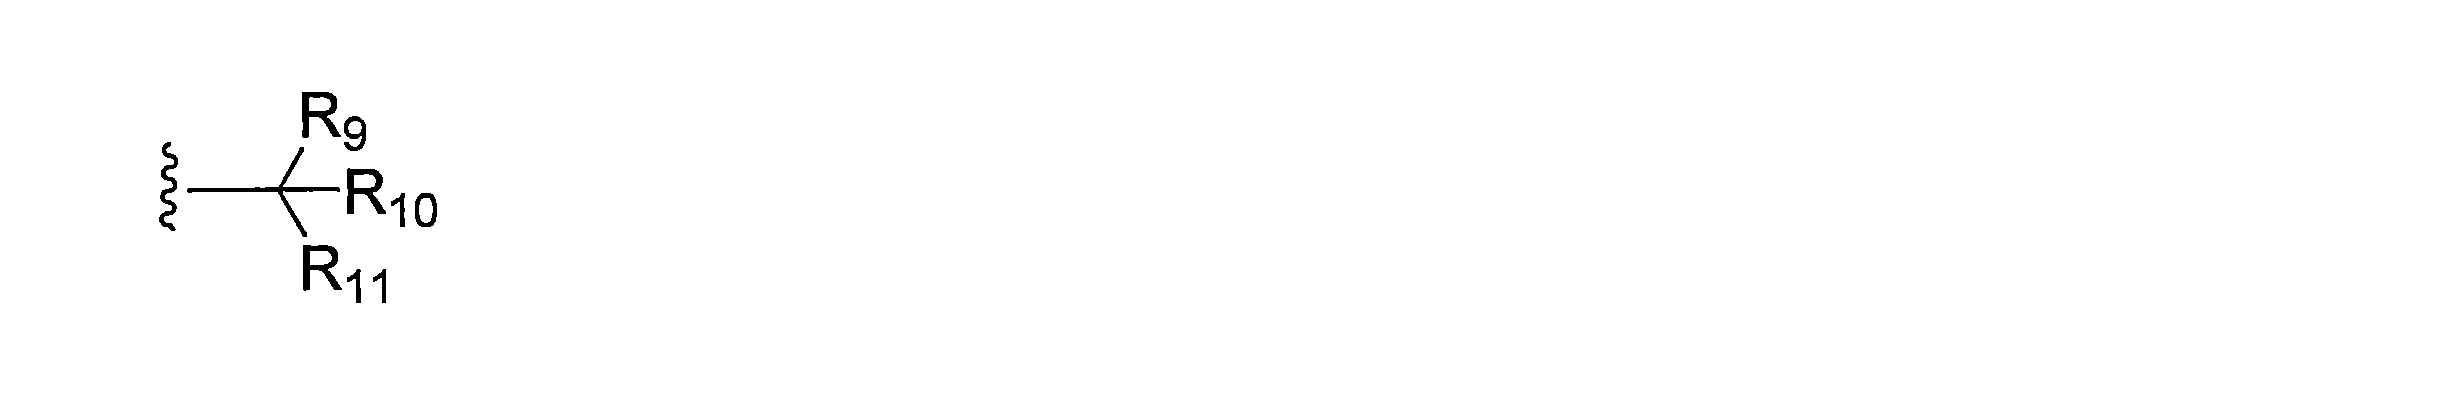
ULM-kのR11は、任意に置換された以下からなる群から選択される。
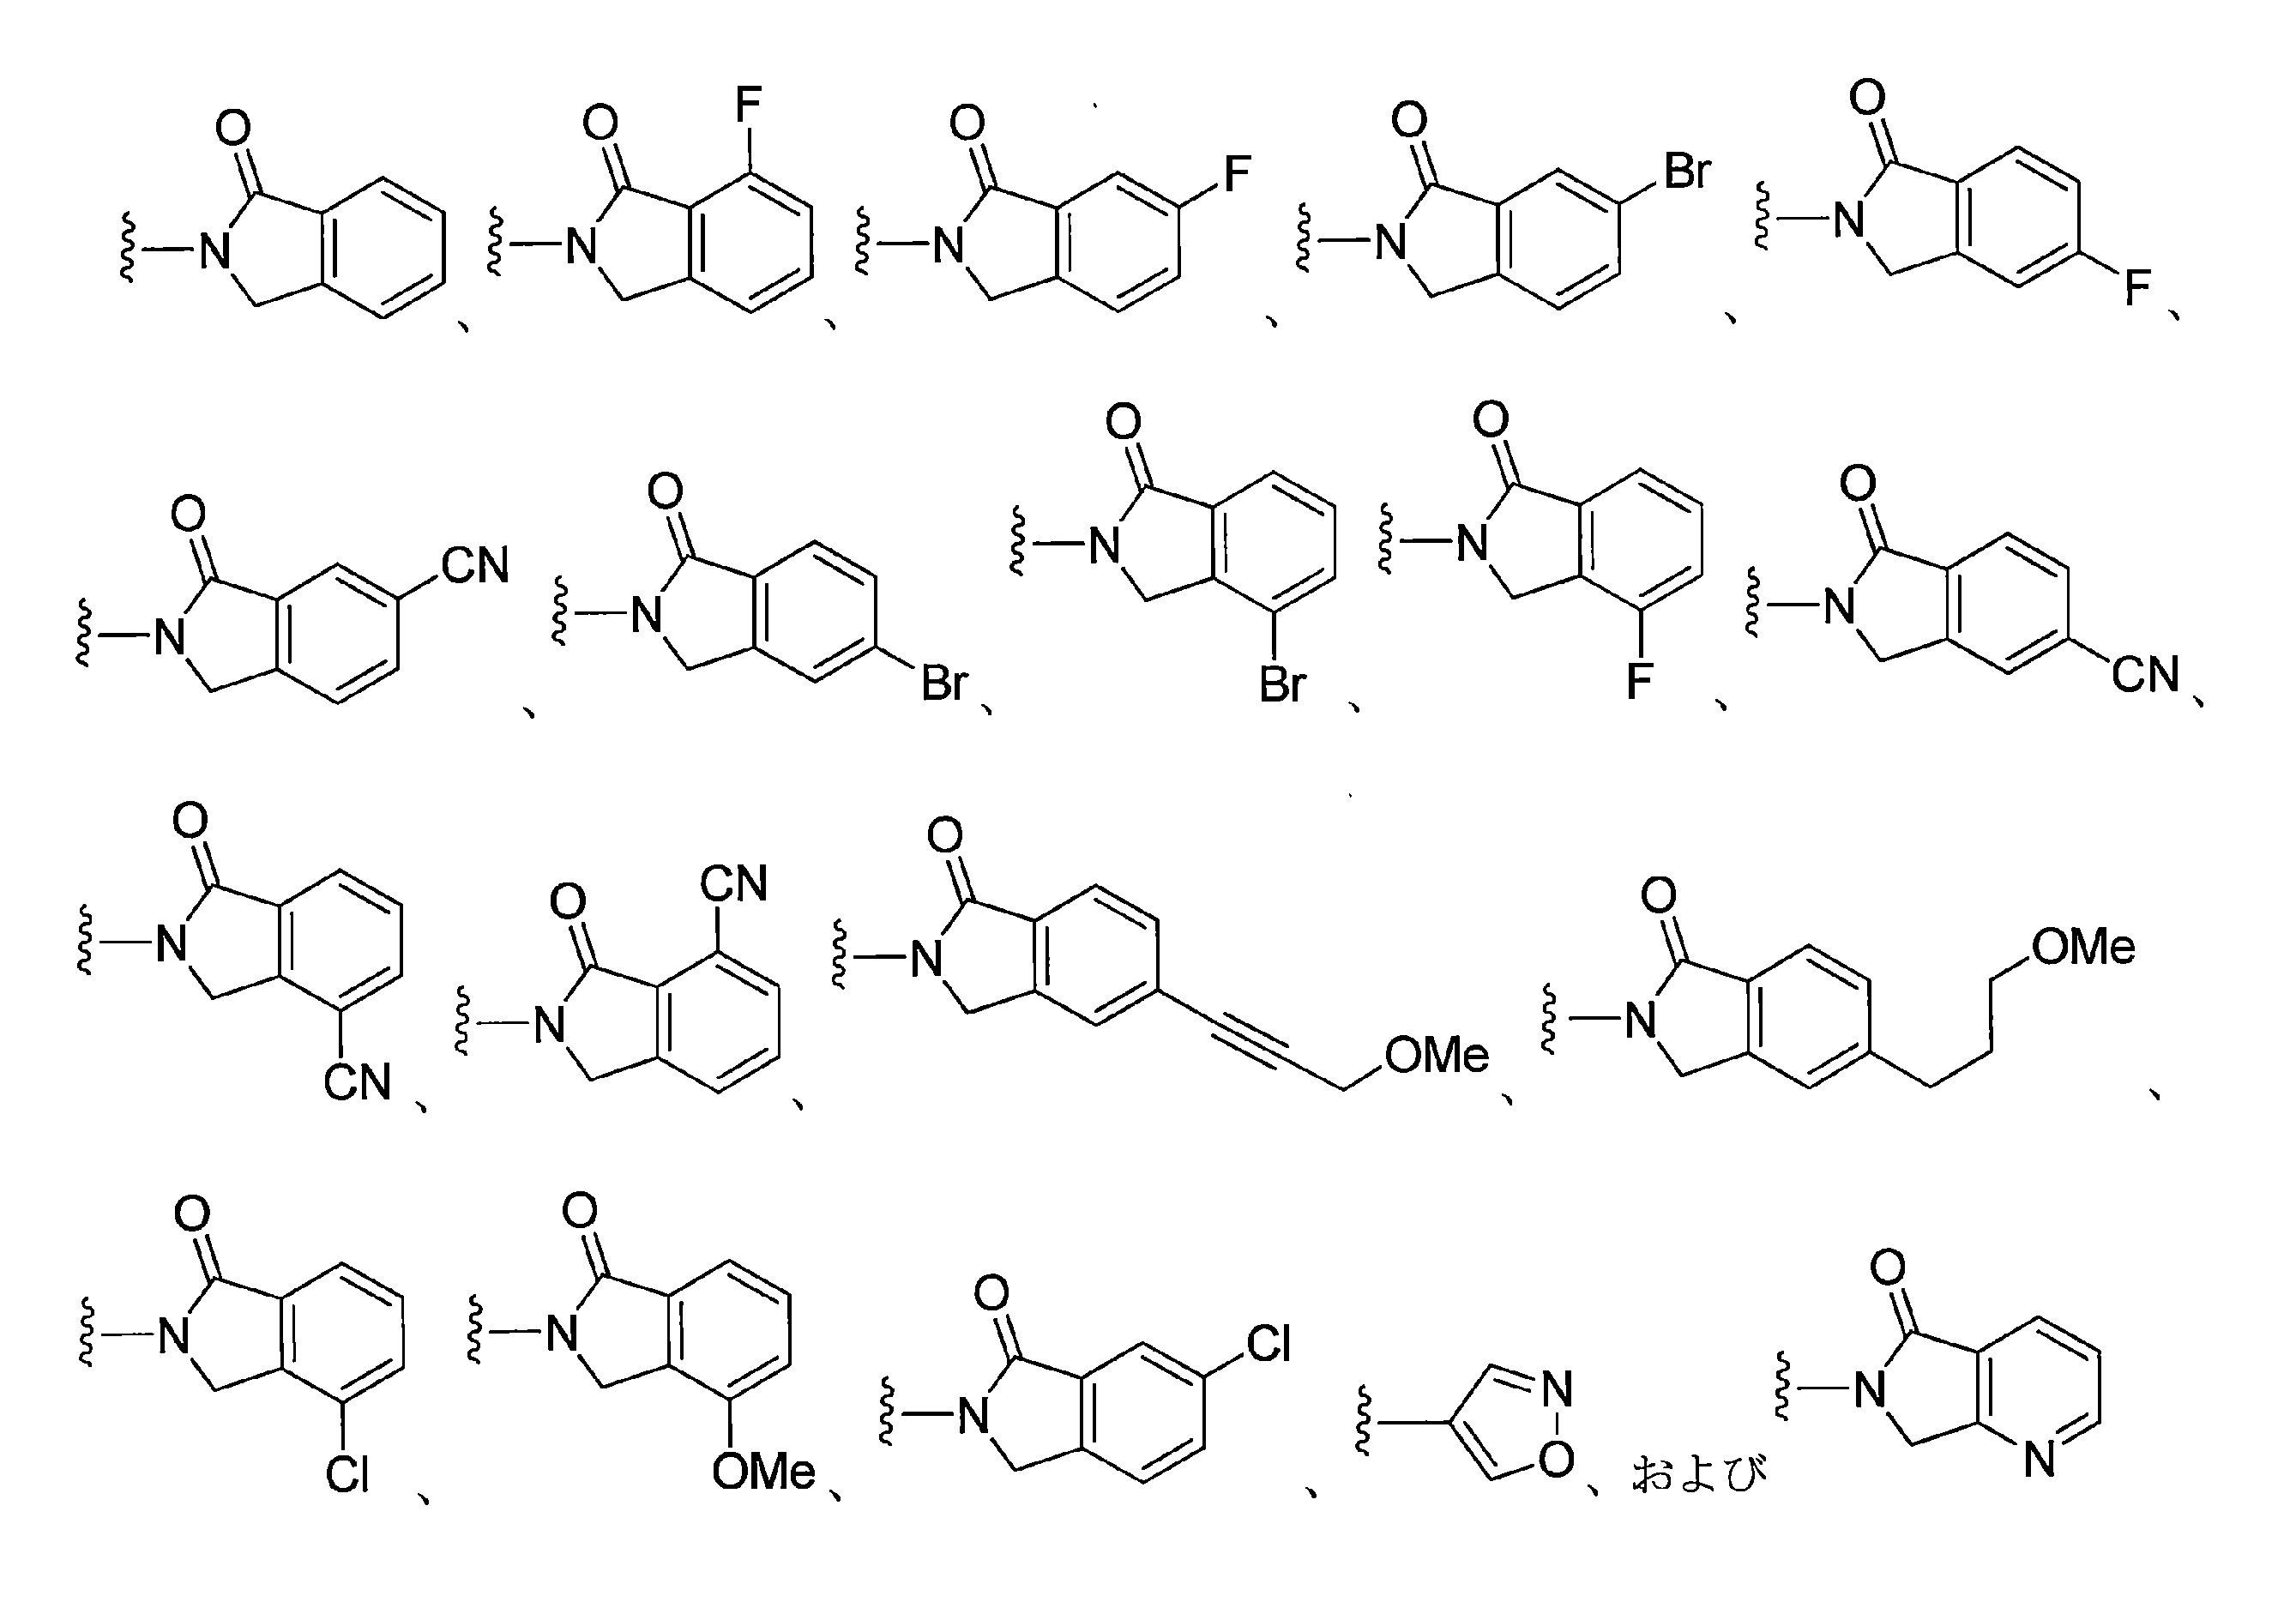
E of ULM-k is C═O,
The M in ULM-k is:
R11 of ULM-k is selected from the group consisting of optionally substituted:
さらに他の実施形態では、以下の化学構造の化合物、

ULM-kのEは、C=Oであり、
ULM-kのR11は、

ULM-kのMは、
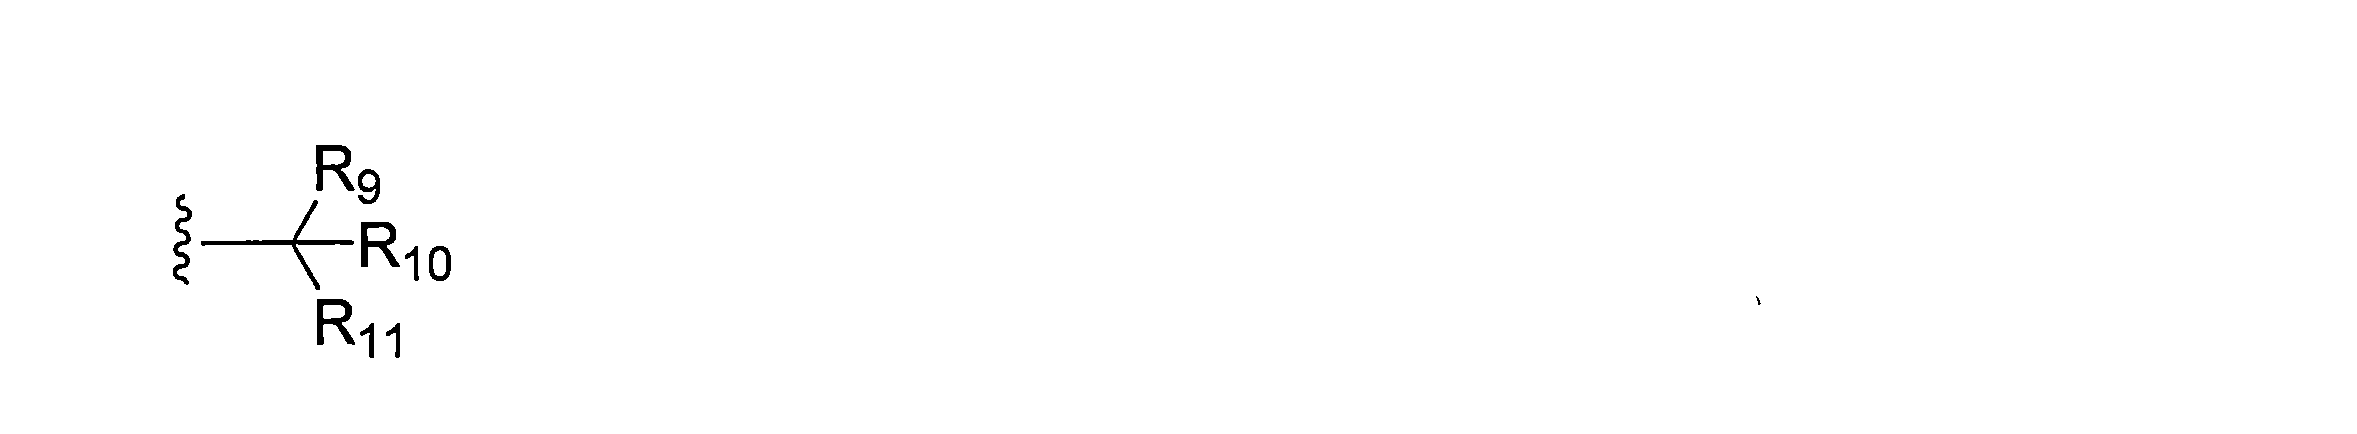
ULM-kのqは、1または2であり、
ULM-kのR20は、H、任意に置換されたアルキル、任意に置換されたシクロアルキル、任意に置換されたアリール、または

ULM-kのR21は、Hまたは任意に置換されたアルキルであり、
ULM-kのR22は、H、任意に置換されたアルキル、任意に置換されたアルコキシ、またはハロアルキルである。
In yet another embodiment, a compound of the following chemical structure:
E of ULM-k is C═O,
R11 of ULM-k is
The M in ULM-k is:
q of ULM-k is 1 or 2;
R20 of ULM-k is H, optionally substituted alkyl, optionally substituted cycloalkyl, optionally substituted aryl, or
R21 of ULM-k is H or optionally substituted alkyl;
R22 of ULM-k is H, optionally substituted alkyl, optionally substituted alkoxy, or haloalkyl.
本明細書に記載される任意の実施形態において、ULM-jまたはULM-kのR11は、以下からなる群から選択される:


特定の実施形態では、ULM-jまたはULM-kのR11は、以下からなる群から選択される:
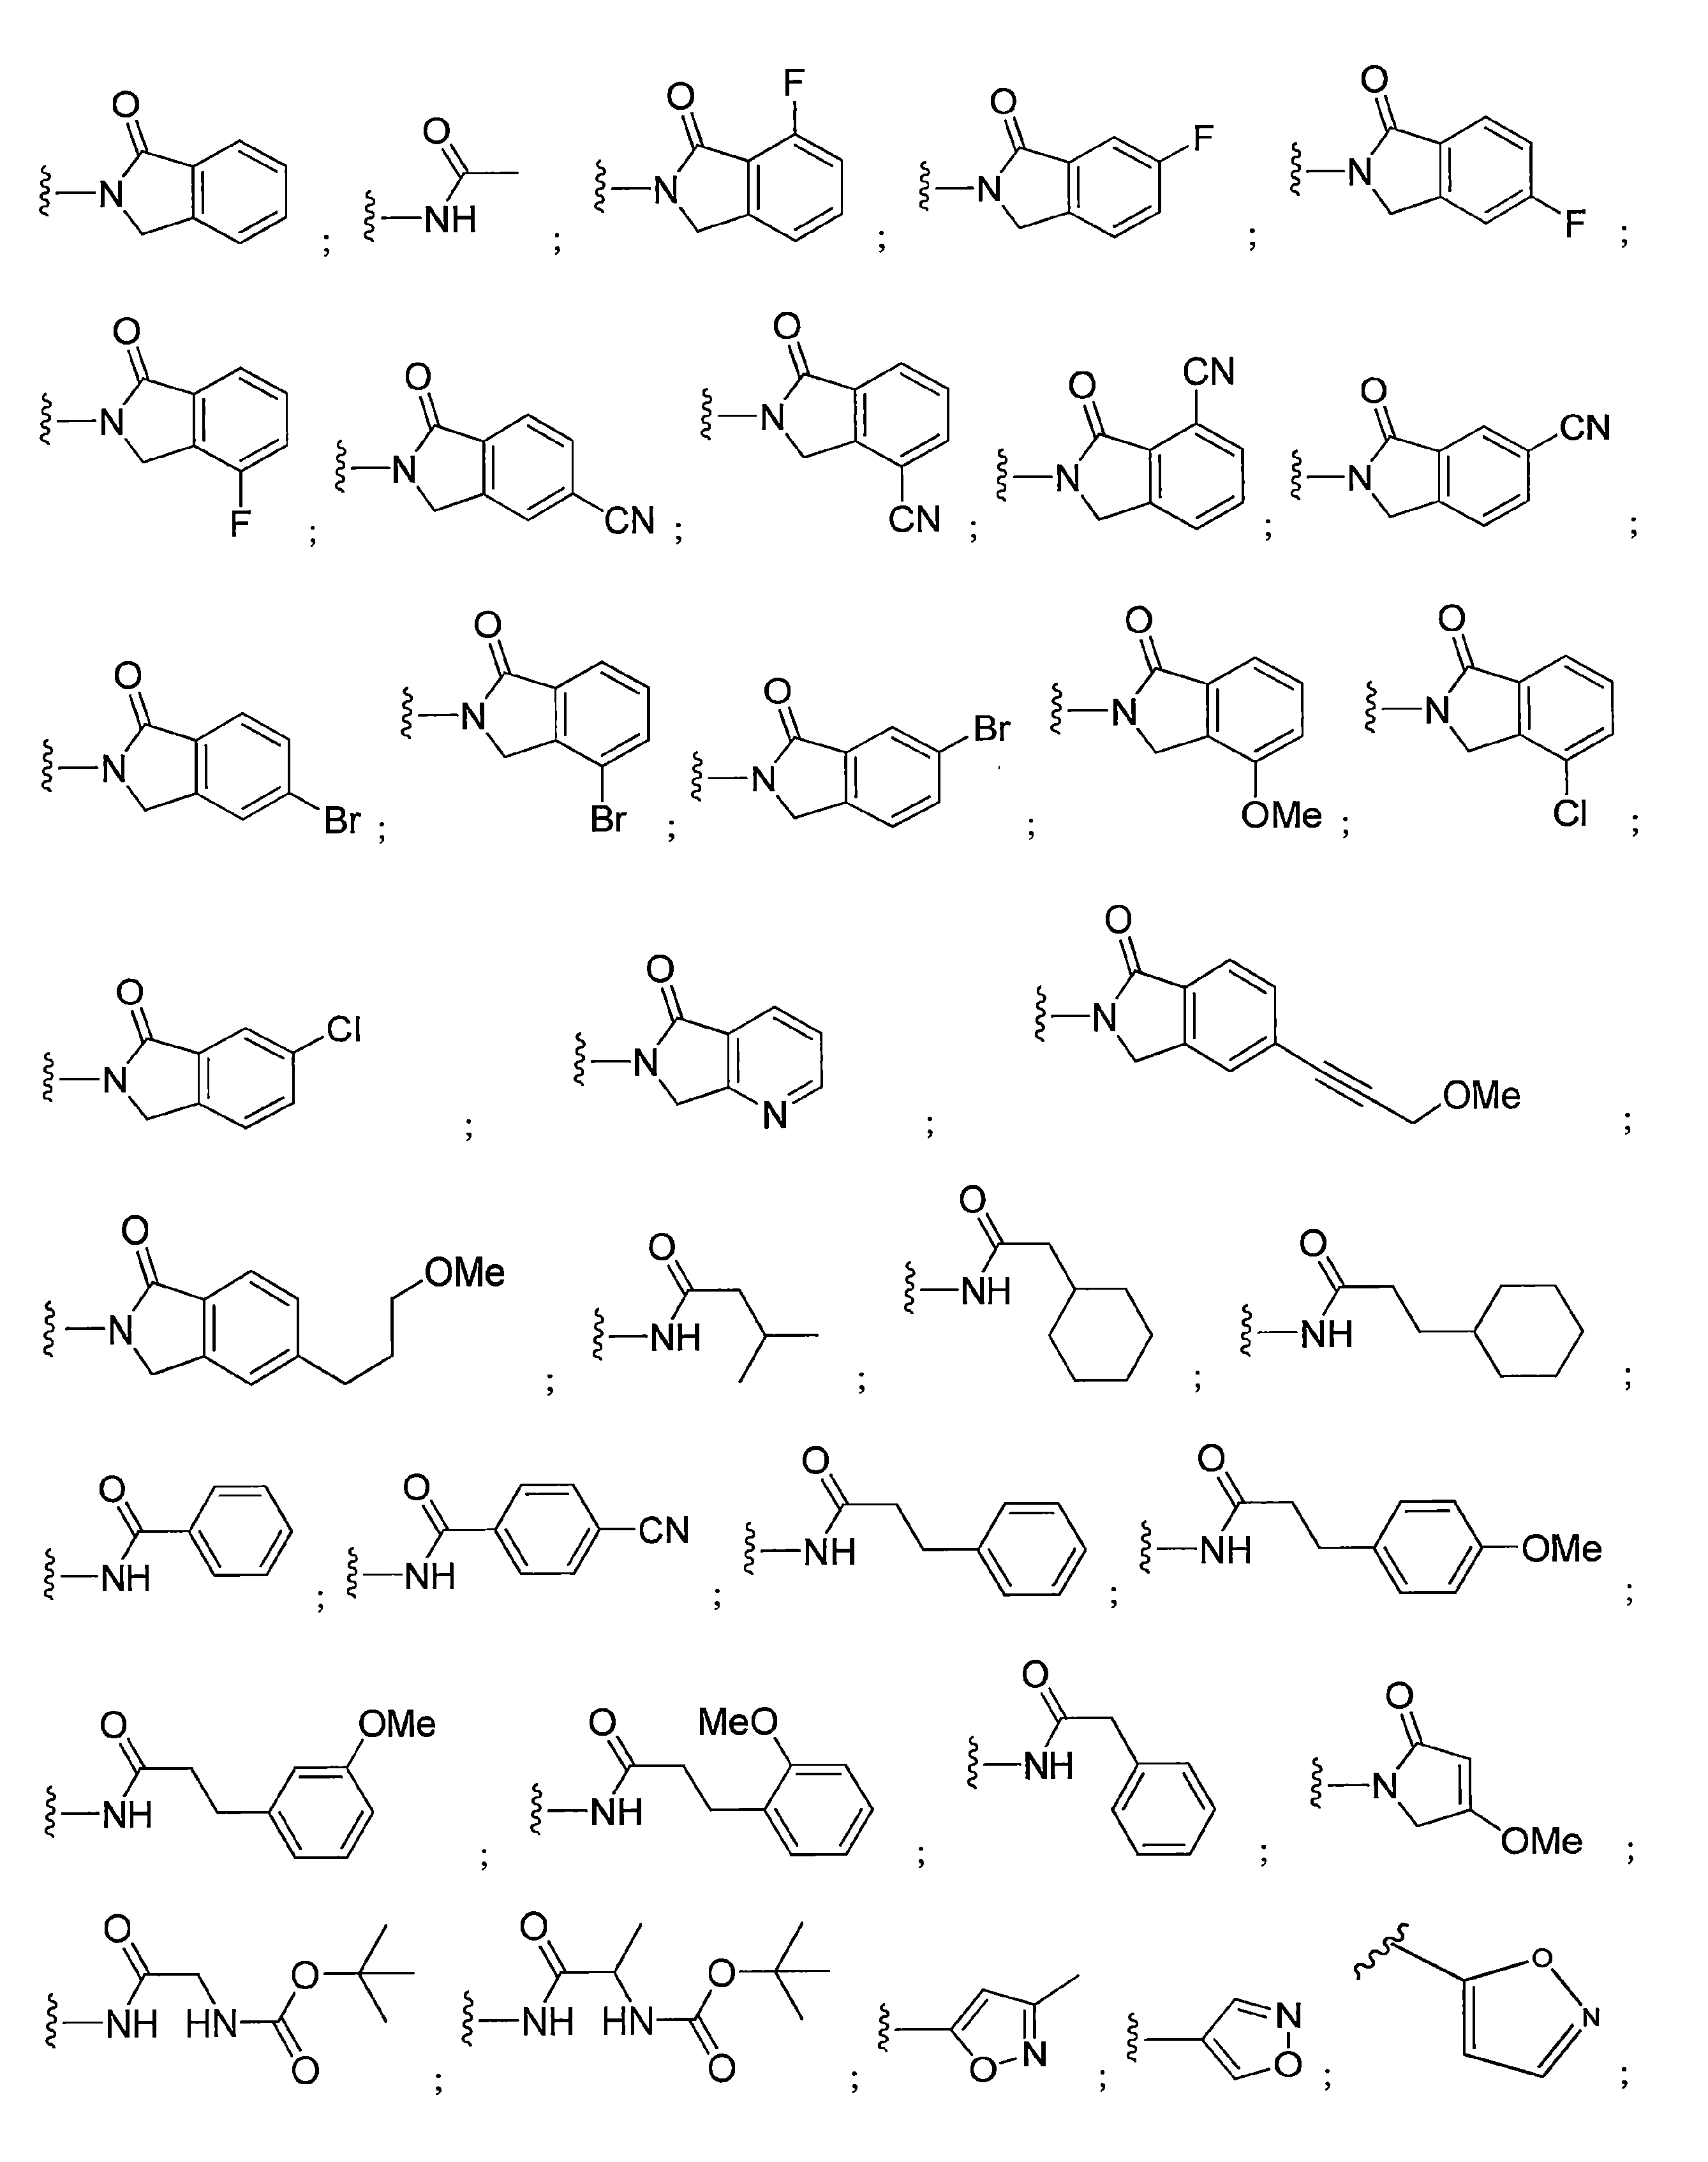

特定の実施形態では、ULM(または存在する場合にはULM’)は、以下の化学構造に従う基である:
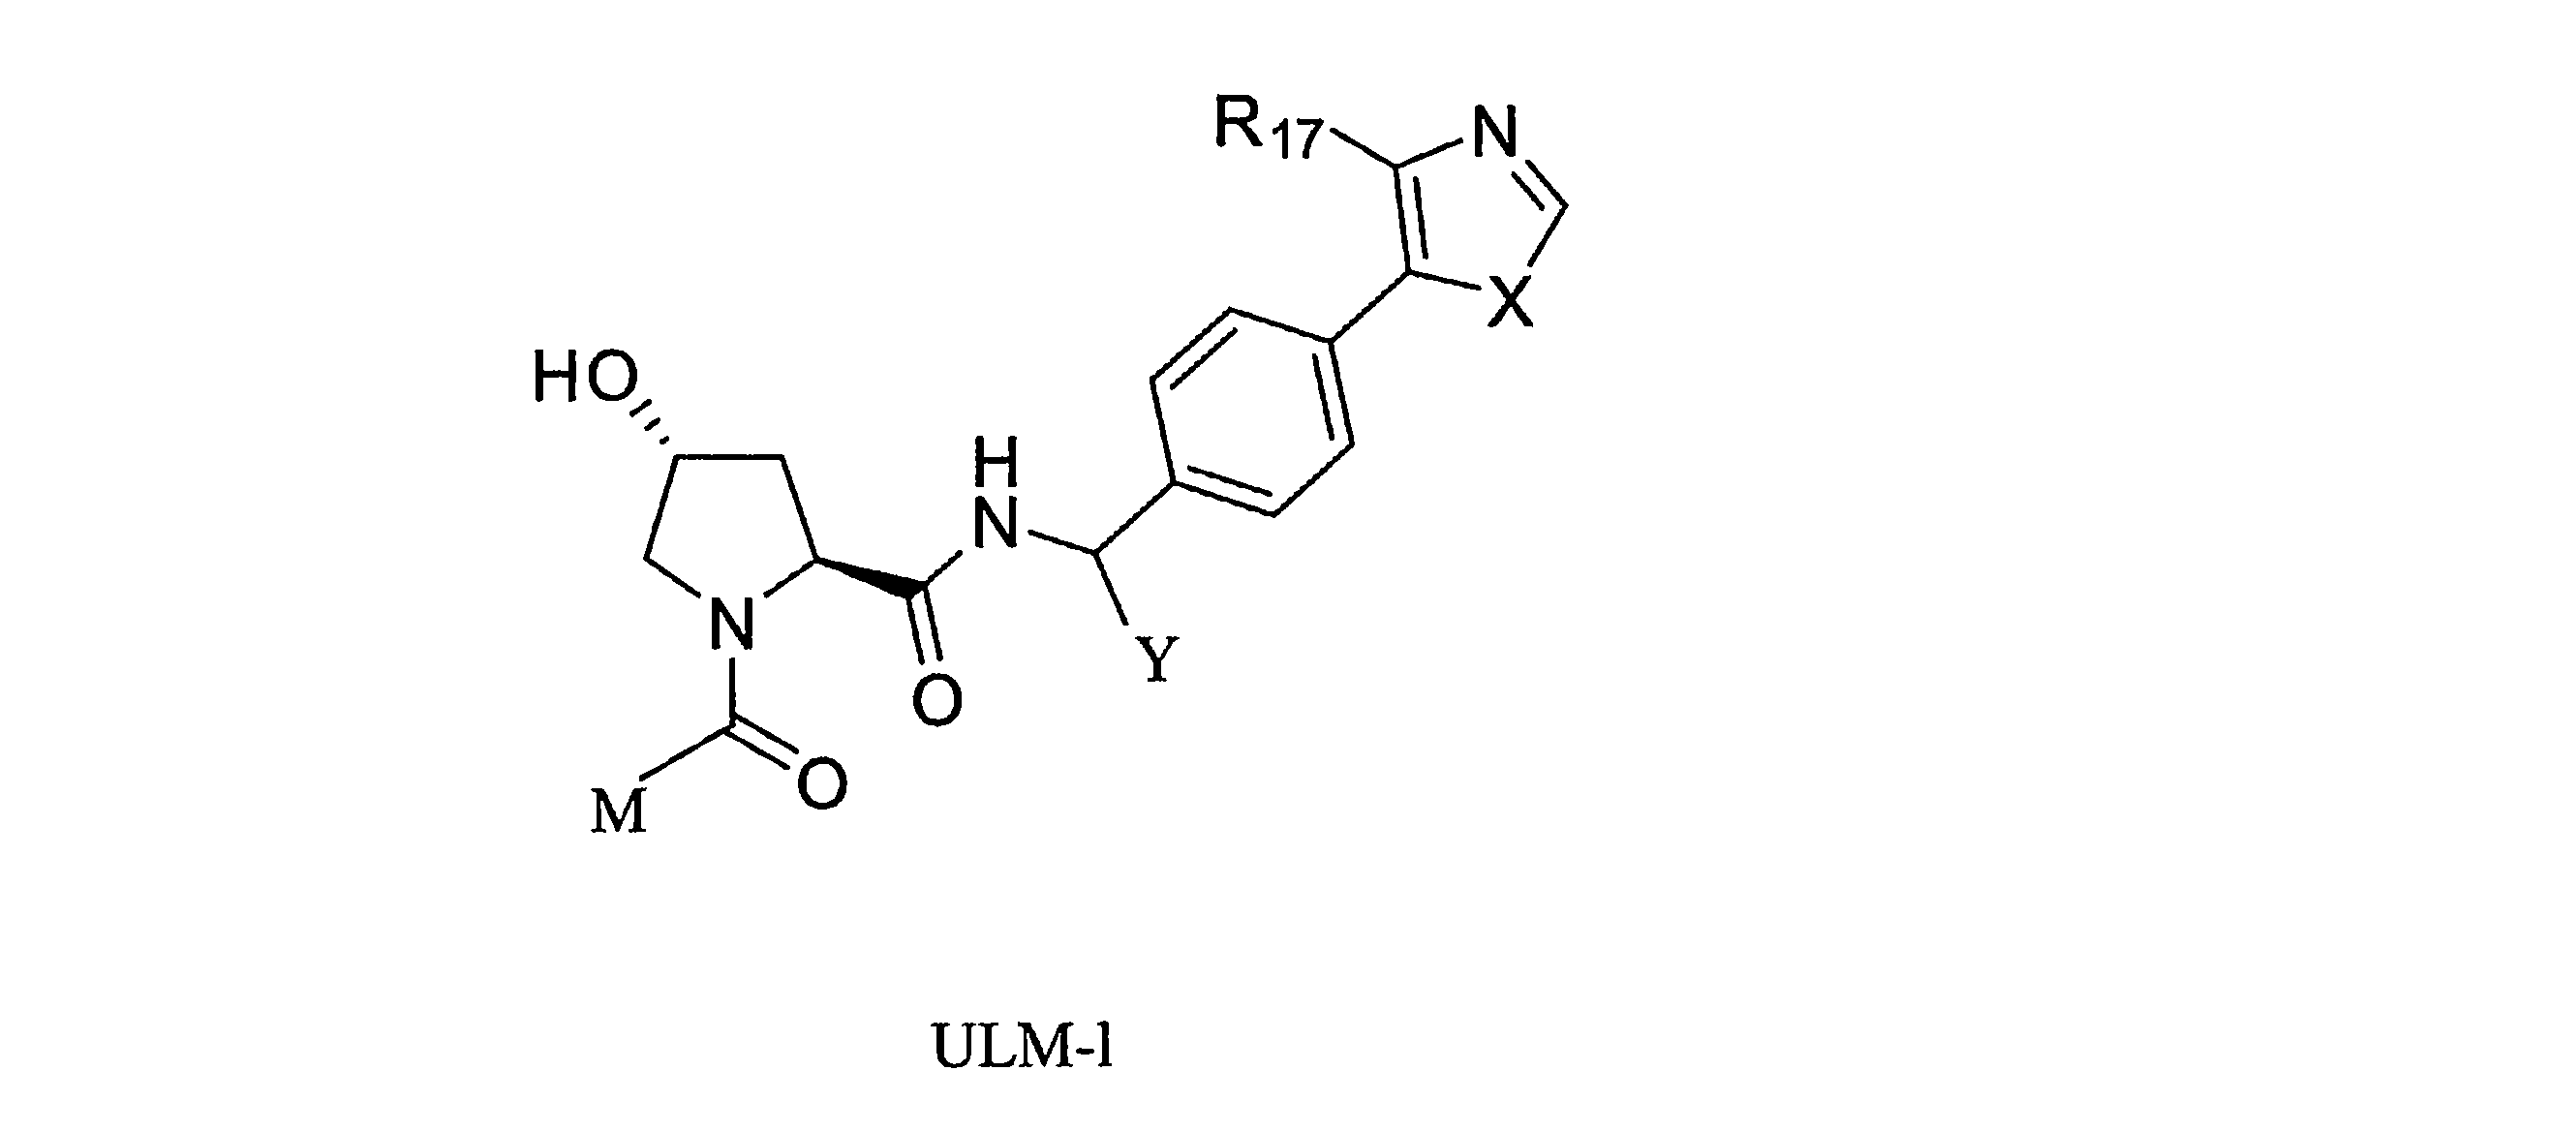
ULM-lのXは、OまたはSであり、
ULM-lのYは、H、メチルまたはエチルであり、
ULM-lのR17は、H、メチル、エチル、ヒドロキシメチル、またはシクロプロピルであり、
ULM-lのMは、任意に置換されたアリール、任意に置換されたヘテロアリール、または
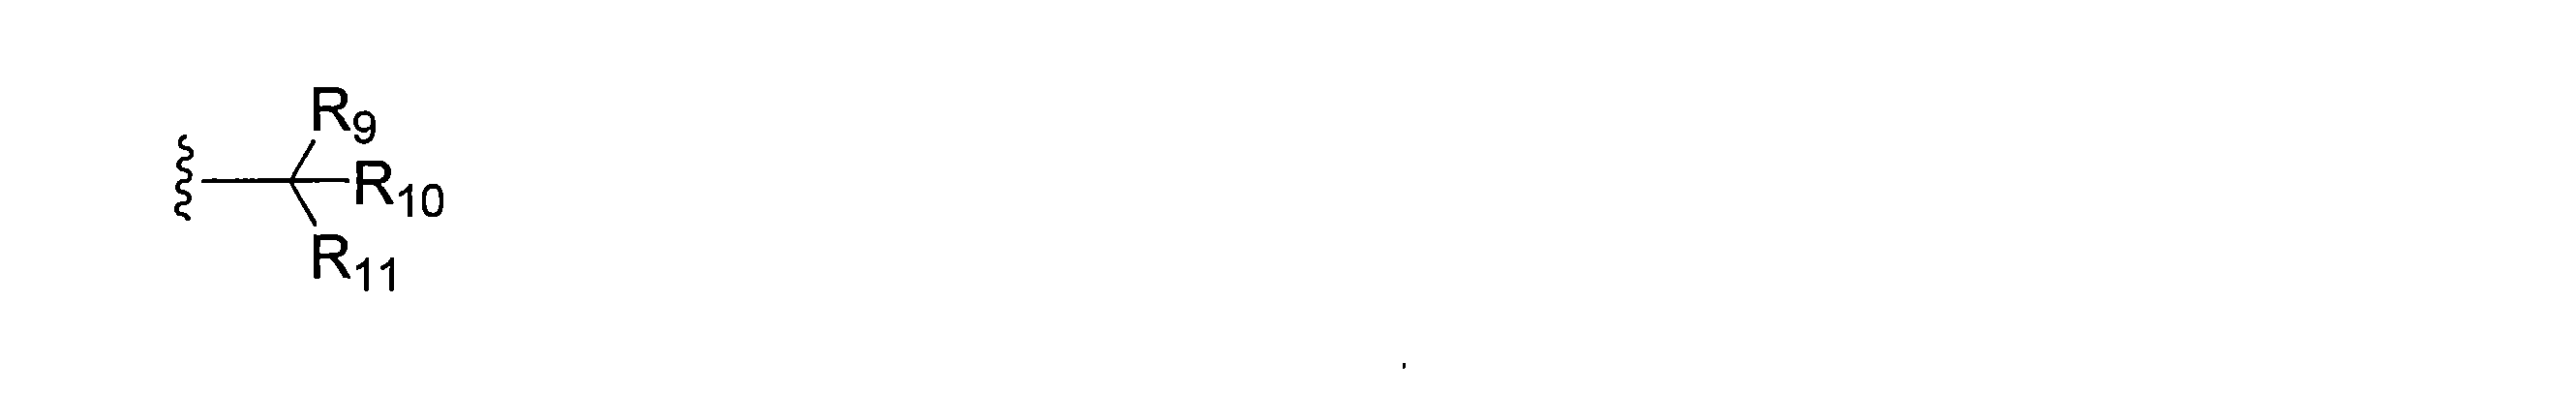
ULM-lのR9は、Hであり、
ULM-lのR10は、H、任意に置換されたアルキル、任意に置換されたハロアルキル、任意に置換されたヘテロアリール、任意に置換されたアリール、任意に置換されたヒドロキシアルキル、任意に置換されたチオアルキル、またはシクロアルキルであり、
ULM-lのR11は、任意に置換された複素芳香族、任意に置換されたヘテロシクリル、任意に置換されたアリール、または

ULM-lのR12は、Hまたは任意に置換されたアルキルであり、
ULM-lのR13は、H、任意に置換されたアルキル、任意に置換されたアルキルカルボニル、任意に置換された(シクロアルキル)アルキルカルボニル、任意に置換されたアラルキルカルボニル、任意に置換されたアリールカルボニル、任意に置換された(ヘテロシクリル)カルボニル、または任意に置換されたアラルキル、任意に置換された(オキソアルキル)カルバメートである。
In certain embodiments, ULM (or ULM′, if present) is a group conforming to the following chemical structure:
X in ULM-1 is O or S;
Y of ULM-1 is H, methyl or ethyl;
R17 of ULM-1 is H, methyl, ethyl, hydroxymethyl, or cyclopropyl;
M of ULM-1 is an optionally substituted aryl, an optionally substituted heteroaryl, or
R9 of ULM-1 is H;
R10 of ULM-1 is H, optionally substituted alkyl, optionally substituted haloalkyl, optionally substituted heteroaryl, optionally substituted aryl, optionally substituted hydroxyalkyl, optionally substituted thioalkyl, or cycloalkyl;
R11 of ULM-1 is an optionally substituted heteroaromatic ring, an optionally substituted heterocyclyl, an optionally substituted aryl, or
R12 of ULM-1 is H or optionally substituted alkyl;
R13 of ULM-1 is H, optionally substituted alkyl, optionally substituted alkylcarbonyl, optionally substituted (cycloalkyl)alkylcarbonyl, optionally substituted aralkylcarbonyl, optionally substituted arylcarbonyl, optionally substituted (heterocyclyl)carbonyl, or optionally substituted aralkyl, optionally substituted (oxoalkyl)carbamate.
一部の実施形態では、ULM、および存在する場合にはULM’はそれぞれ独立して、以下の化学構造に従う基である:

ULM-mのYは、H、メチオール(methyol)またはエチルである
ULM-mのR9は、Hであり、
R10は、イソプロピル、tert-ブチル、sec-ブチル、シクロペンチル、またはシクロヘキシルであり、
ULM-mのR11は、任意に置換されたアミド、任意に置換されたイソインドリノン、任意に置換されたイソキサゾール、任意に置換されたヘテロシクリルである。
In some embodiments, ULM, and, if present, ULM′, are each independently a group conforming to the following chemical structure:
Y of ULM-m is H, methyol or ethyl; R9 of ULM-m is H;
R10 is isopropyl, tert-butyl, sec-butyl, cyclopentyl, or cyclohexyl;
R11 of ULM-m is an optionally substituted amide, an optionally substituted isoindolinone, an optionally substituted isoxazole, or an optionally substituted heterocyclyl.
本開示の他の好ましい実施形態では、ULM、および存在する場合ULM’は、各々独立して、以下の化学構造に従う基であり、

ULM-nのR17は、メチル、エチル、またはシクロプロピルであり、
ULM-nのR9、R10、およびR11は、上記に定義されるとおりである。他の事例では、R9は、Hであり、
ULM-nのR10は、H、アルキル、またはシクロアルキル(好ましくはイソプロピル、tert-ブチル、sec-ブチル、シクロペンチル、またはシクロヘキシル)である。
In another preferred embodiment of the present disclosure, ULM, and, if present, ULM′, are each independently a group according to the following chemical structure:
R17 of ULM-n is methyl, ethyl, or cyclopropyl;
R9, R10, and R11 of ULM-n are as defined above. In other instances, R9 is H;
R10 of ULM-n is H, alkyl, or cycloalkyl (preferably isopropyl, tert-butyl, sec-butyl, cyclopentyl, or cyclohexyl).
本開示の他の好ましい実施形態では、ULM、および存在する場合ULM’は、各々独立して、以下の化学構造に従う基であり、
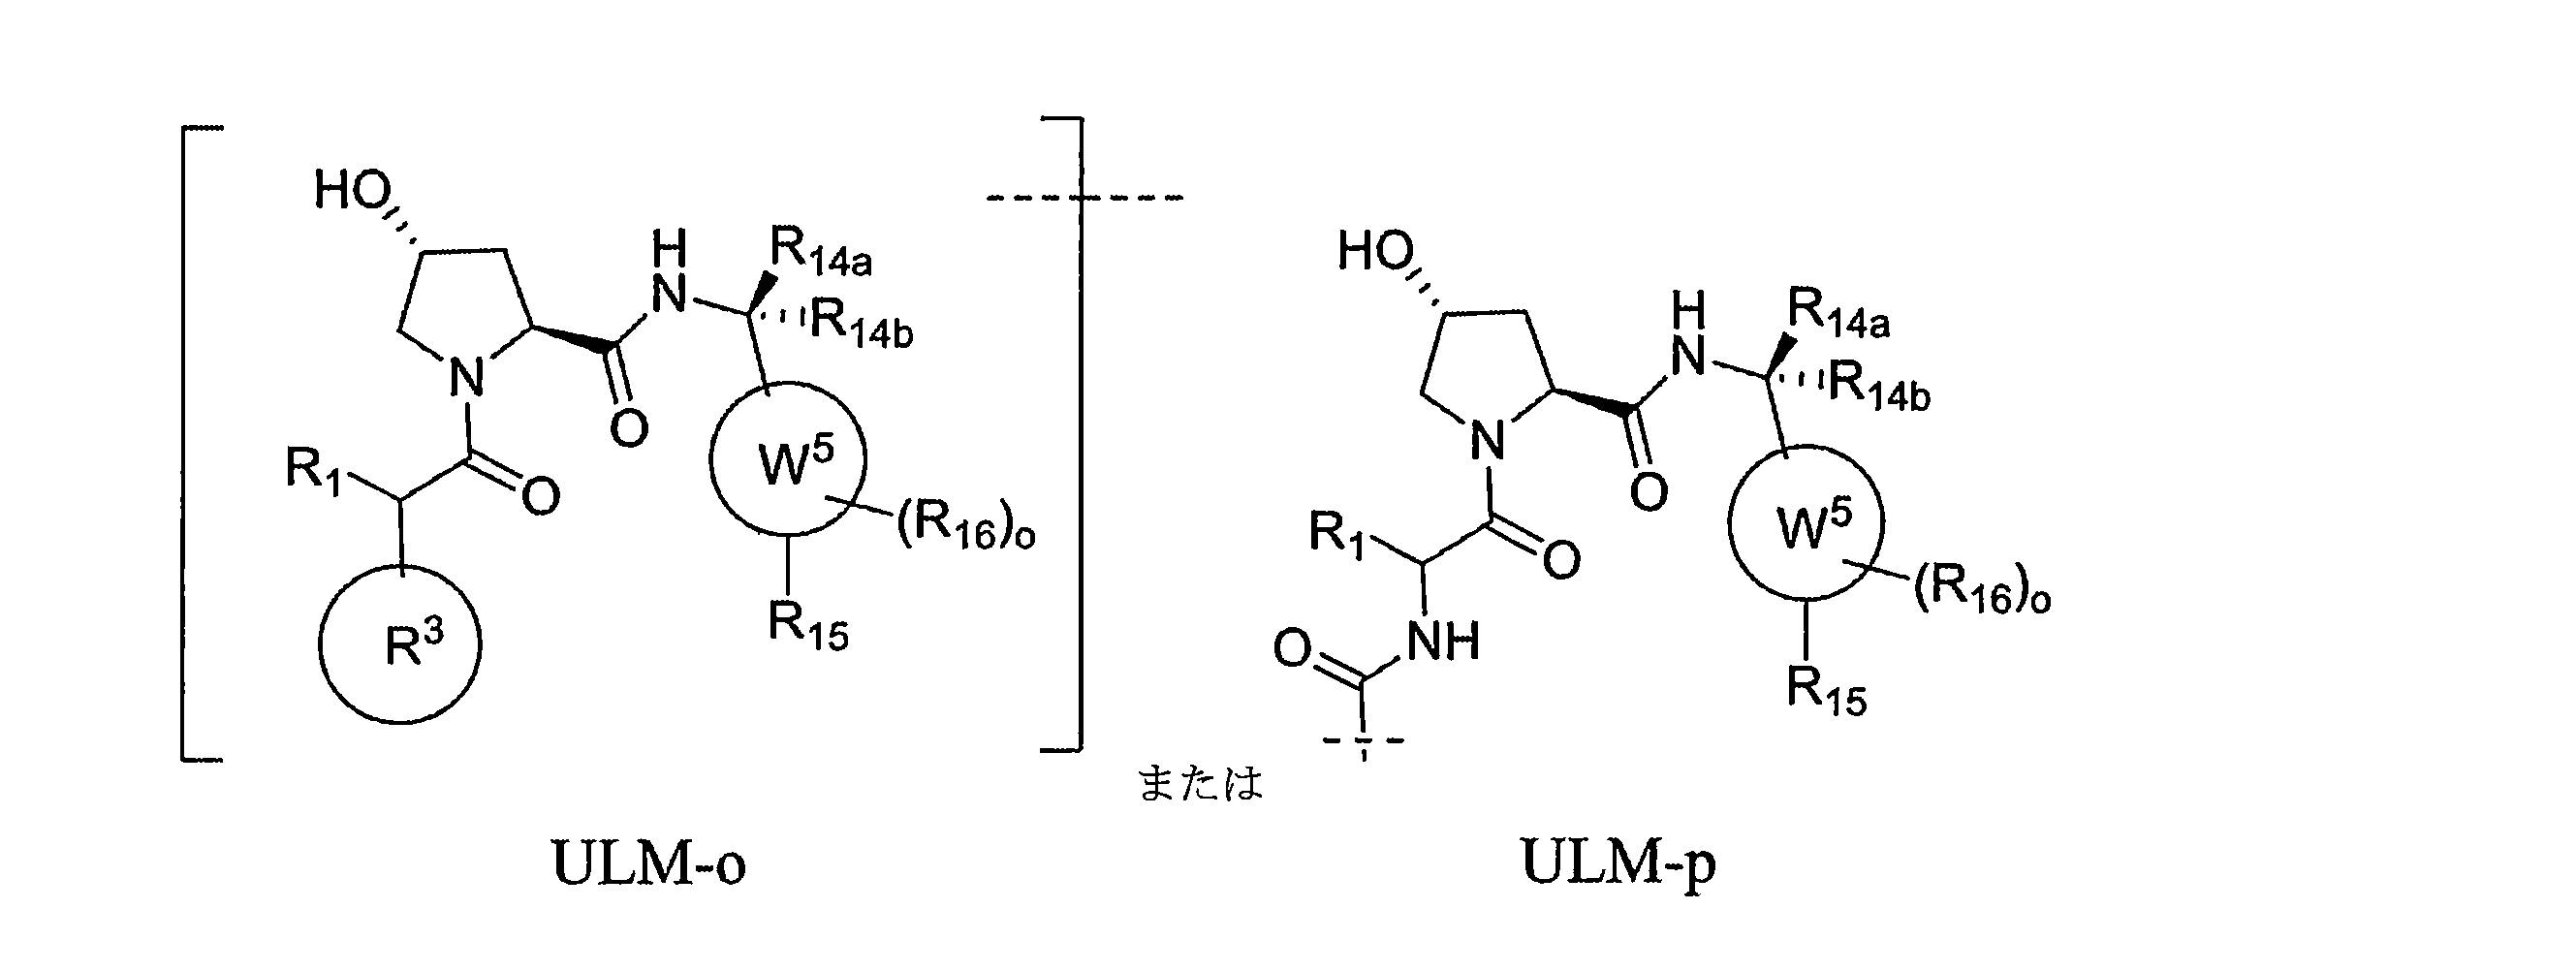
R1は、H、任意に置換されたアルキル、または任意に置換されたシクロアルキルであり、
R3は、任意に置換された5~6員ヘテロアリールであり、
W5は、任意に置換されたフェニル、任意に置換されたナプチル、または任意に置換されたピリジニルであり、
R14aおよびR14bのうちの一方は、H、任意に置換されたアルキル、任意に置換されたハロアルキル(例えば、フルオロアルキル)、任意に置換されたアルコキシ、任意に置換されたヒドロキシルアルキル、任意に置換されたアルキルアミン、任意に置換されたヘテロルキル、任意に置換されたアルキル-ヘテロシクロアルキル、任意に置換されたアルコキシ-ヘテロシクロアルキル、COR26、CONR27aR27b、NHCOR26、もしくはNHCH3COR26であり、R14aおよびR14bのうちの他方は、Hであるか、またはR14a、R14bは、それらが結合している炭素原子と一緒になって、任意に置換された3~6員シクロアルキル、ヘテロシクロアルキル、スピロシクロアルキル、もしくはスピロヘテロシクリルを形成し、スピロヘテロシクリルは、エポキシドまたはアジリジンではなく、
R15は、CN、任意に置換されたフルオロアルキル、


各R16は、独立して、ハロ、CN、任意に置換されたアルキル、任意に置換されたハロアルキル、ヒドロキシ、またはハロアルコキシから選択され、
各R26は、独立して、H、任意に置換されたアルキル、またはNR27aR27bであり、
各R27aおよびR27bは、独立して、H、任意に置換されたアルキル、任意に置換されたシクロアルキル(例えば、任意に置換された3~5員シクロアルキル)であるか、またはR27aおよびR27bは、それらが結合している窒素原子と一緒になって、4~6員ヘテロシクリルを形成し、
各R28は、独立して、H、ハロゲン、CN、任意に置換されたアミノアルキル、任意に置換されたアミドアルキル、任意に置換されたハロアルキル、任意に置換されたアルキル、任意に置換されたアルコキシ、任意に置換されたヘテロアルキル、任意に置換されたアルキルアミン、任意に置換されたヒドロキシアルキル、アミン、任意に置換されたアルキニル、または任意に置換されたシクロアルキルであり、
oは、0、1、または2であり、
pは、0、1、2、3、または4である。
In another preferred embodiment of the present disclosure, ULM, and, if present, ULM′, are each independently a group according to the following chemical structure:
R1 is H, optionally substituted alkyl, or optionally substituted cycloalkyl;
R3 is an optionally substituted 5-6 membered heteroaryl;
W5 is optionally substituted phenyl, optionally substituted naphthyl, or optionally substituted pyridinyl;
one of R14a and R14b is H, optionally substituted alkyl, optionally substituted haloalkyl (e.g., fluoroalkyl), optionally substituted alkoxy, optionally substituted hydroxylalkyl, optionally substituted alkylamine, optionally substituted heteroalkyl, optionally substituted alkyl-heterocycloalkyl, optionally substituted alkoxy-heterocycloalkyl, COR26, CONR27aR27b, NHCOR26, or NHCH3COR26; the other of R14a and R14b is H, or R14a, R14b together with the carbon atom to which they are attached form an optionally substituted 3- to 6-membered cycloalkyl, heterocycloalkyl, spirocycloalkyl, or spiroheterocyclyl, wherein the spiroheterocyclyl is not an epoxide or aziridine;
R15 is CN, optionally substituted fluoroalkyl,
each R16 is independently selected from halo, CN, optionally substituted alkyl, optionally substituted haloalkyl, hydroxy, or haloalkoxy;
each R26 is independently H, optionally substituted alkyl, or NR27aR27b;
each R27a and R27b is independently H, optionally substituted alkyl, optionally substituted cycloalkyl (e.g., optionally substituted 3- to 5-membered cycloalkyl), or R27a and R27b together with the nitrogen atom to which they are attached form a 4- to 6-membered heterocyclyl;
each R28 is independently H, halogen, CN, optionally substituted aminoalkyl, optionally substituted amidoalkyl, optionally substituted haloalkyl, optionally substituted alkyl, optionally substituted alkoxy, optionally substituted heteroalkyl, optionally substituted alkylamine, optionally substituted hydroxyalkyl, amine, optionally substituted alkynyl, or optionally substituted cycloalkyl;
o is 0, 1, or 2;
p is 0, 1, 2, 3, or 4.
本明細書に記載される態様または実施形態のいずれにおいても、ULMは、以下の式:

X4、X5、およびX6の各々が、CHおよびNから選択され、2個以下が、Nであり、
R1は、C1-6アルキルであり、
R3は、ULM-oおよびULM-pについて定義されたものと同じであり、
R14aおよびR14bのうちの一方は、H、任意に置換されたアルキル、任意に置換されたハロアルキル、任意に置換されたアルコキシ、任意に置換されたヒドロキシルアルキル、任意に置換されたアルキルアミン、任意に置換されたアミド、任意に置換されたアルキル-アミド、任意に置換されたアルキル-シアノ、任意に置換されたアルキル-ホスフェート、任意に置換されたヘテロアルキル、任意に置換されたアルキル-ヘテロシクロアルキル、任意に置換されたアルコキシ-ヘテロシクロアルキル、COR26、CONR27aR27b、NHCOR26、もしくはNHCH3COR26であり、R14aおよびR14bのうちの他方が、Hであるか、またはR14aおよびR14bは、それらが結合している炭素原子と一緒になって、任意に置換された3~5員シクロアルキル、ヘテロシクロアルキル、スピロシクロアルキル、もしくはスピロヘテロシクリルを形成し、スピロヘテロシクリルは、エポキシドまたはアジリジンではなく、
各R27aおよびR27bは、独立して、H C1-6アルキルまたはシクロルキル(例えば、任意に置換された3~5員シクロアルキル)であり、
oは、0、1、または2であり、
qは、1、2、3、または4であり、
R15は、任意に置換された

R28は、H、メチル、CH2N(Me)2、CH2OH、CH2O(C1-4アルキル)、CH2NHC(O)C1-4アルキル、NH2、
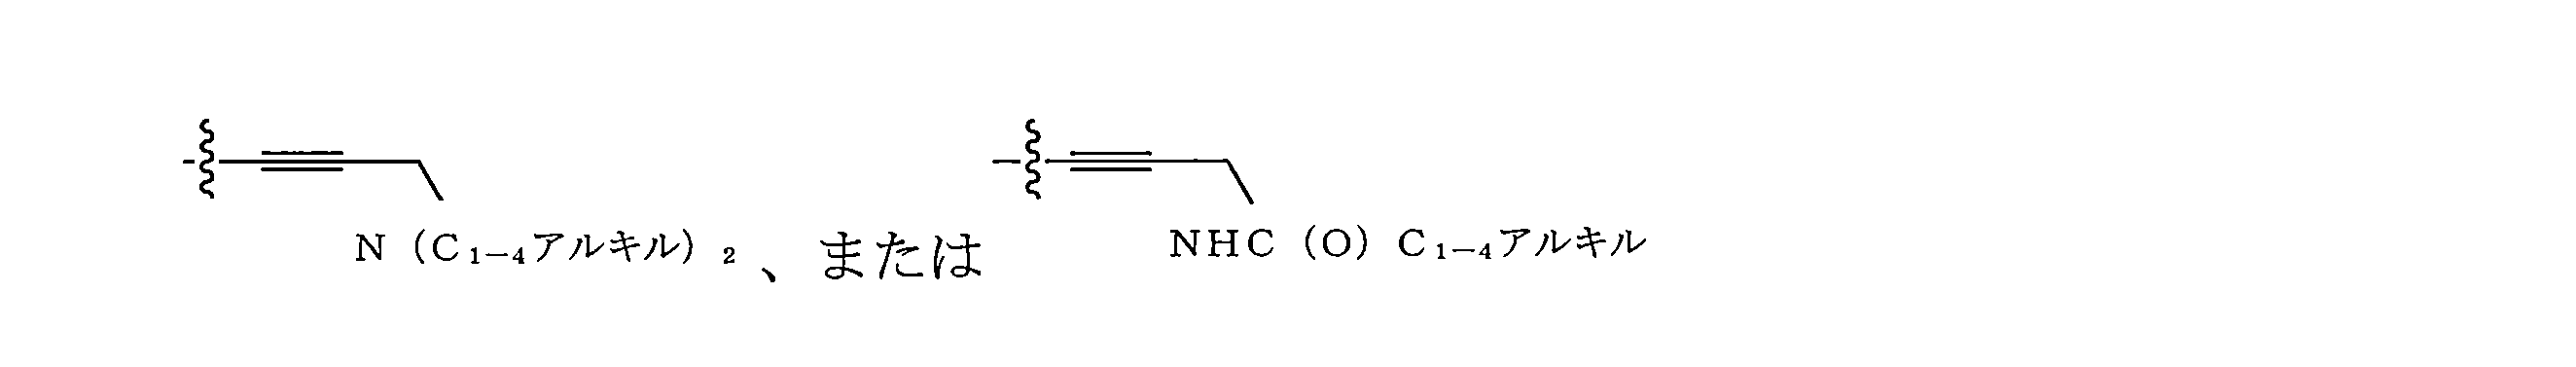
R28Cは、H、メチル、フルオロ、またはクロロであり、
R16は、H、C1-4アルキル、フルオロ、クロロ、CN、またはC1-4アルコキシである。
In any of the aspects or embodiments described herein, the ULM has the following formula:
Each of X4, X5, and X6 is selected from CH and N, with up to 2 being N;
R1 is C1-6 alkyl;
R3 is the same as defined for ULM-o and ULM-p;
one of R14a and R14b is H, optionally substituted alkyl, optionally substituted haloalkyl, optionally substituted alkoxy, optionally substituted hydroxylalkyl, optionally substituted alkylamine, optionally substituted amide, optionally substituted alkyl-amide, optionally substituted alkyl-cyano, optionally substituted alkyl-phosphate, optionally substituted heteroalkyl, optionally substituted alkyl-heterocycloalkyl, optionally substituted alkoxy-heterocycloalkyl, COR26, CONR27aR27b, NHCOR26, or NHCH3COR26; the other of R14a and R14b is H; or R14a and R14b together with the carbon atom to which they are attached form an optionally substituted 3- to 5-membered cycloalkyl, heterocycloalkyl, spirocycloalkyl, or spiroheterocyclyl, wherein the spiroheterocyclyl is not an epoxide or aziridine;
each R27a and R27b is independently H C1-6 alkyl or cycloalkyl (e.g., optionally substituted 3- to 5-membered cycloalkyl);
o is 0, 1, or 2;
q is 1, 2, 3, or 4;
R15 is optionally substituted
R28 is H, methyl, CH2N(Me)2, CH2OH, CH2O(C1-4 alkyl), CH2NHC(O)C1-4 alkyl, NH2,
R28C is H, methyl, fluoro, or chloro;
R16 is H, C1-4 alkyl, fluoro, chloro, CN, or C1-4 alkoxy.
本明細書に記載される任意の態様または実施形態では、R14aおよびR14bは、H、C1-4アルキル、C1-4シクロアルキル、C1-4ハロアルキル、C1-4ヒドロキシアルキル、C1-4アルキルオキシアルキル、C1-4アルキル-NR27aR27b、およびCONR27aR27bから選択される。 In any aspect or embodiment described herein, R14a and R14b are selected from H, C1-4 alkyl, C1-4 cycloalkyl, C1-4 haloalkyl, C1-4 hydroxyalkyl, C1-4 alkyloxyalkyl, C1-4 alkyl-NR27aR27b, and CONR27aR27b.
本明細書に記載される任意の態様または実施形態では、R14aおよびR14bのうちの少なくとも一方は、Hである(例えば、R14aおよびR14bの両方は、Hである)。 In any aspect or embodiment described herein, at least one of R14a and R14b is H (e.g., both R14a and R14b are H).
本明細書に記載される任意の態様または実施形態では、R14aおよびR14bのうちの少なくとも一方は、任意に置換されたアルキル、任意に置換されたハロアルキル、任意に置換されたアルコキシ、任意に置換されたヒドロキシルアルキル、任意に置換されたアルキルアミン、任意に置換されたヘテロルキル、任意に置換されたアルキル-ヘテロシクロアルキル、任意に置換されたアルコキシ-ヘテロシクロアルキル、COR26、CONR27aR27b、NHCOR26、またはNHCH3COR26である。あるいは、本明細書に記載される任意の態様または実施形態では、R14aおよびR14bのうちの一方は、任意に置換されたアルキル、任意に置換されたハロアルキル、任意に置換されたアルコキシ、任意に置換されたヒドロキシルアルキル、任意に置換されたアルキルアミン、任意に置換されたヘテロルキル、任意に置換されたアルキルヘテロシクロアルキル、任意に置換されたアルコキシヘテロシクロアルキル、COR26、CONR27aR27b、NHCOR26、またはNHCH3COR26であり、R14aおよびR14bのうちの他方は、Hである。 In any aspect or embodiment described herein, at least one of R14a and R14b is an optionally substituted alkyl, an optionally substituted haloalkyl, an optionally substituted alkoxy, an optionally substituted hydroxylalkyl, an optionally substituted alkylamine, an optionally substituted heteroalkyl, an optionally substituted alkyl-heterocycloalkyl, an optionally substituted alkoxy-heterocycloalkyl, COR26, CONR27aR27b, NHCOR26, or NHCH3COR26. Alternatively, in any aspect or embodiment described herein, one of R14a and R14b is optionally substituted alkyl, optionally substituted haloalkyl, optionally substituted alkoxy, optionally substituted hydroxylalkyl, optionally substituted alkylamine, optionally substituted heteroalkyl, optionally substituted alkylheterocycloalkyl, optionally substituted alkoxyheterocycloalkyl, COR26, CONR27aR27b, NHCOR26, or NHCH3COR26, and the other of R14a and R14b is H.
本明細書に記載される任意の態様または実施形態では、R14aおよびR14bは、それらが結合している炭素原子と一緒になって、

本開示の他の好ましい実施形態では、ULM、および存在する場合ULM’は、各々独立して、以下の化学構造に従う基であり、

Xは、CHまたはNであり、
ULM-qおよびULM-rのR1、R3、R14a、R14b、およびR15が、ULM-oおよびULM-pについて定義されたものと同じである。
In other preferred embodiments of the present disclosure, ULM, and, if present, ULM′, are each independently a group according to the following chemical structure:
X is CH or N;
R1, R3, R14a, R14b, and R15 of ULM-q and ULM-r are the same as defined for ULM-o and ULM-p.
本明細書に記載される態様または実施形態のいずれかにおいて、本明細書に記載されるULM(または存在する場合、ULM’)は、その薬学的に許容可能な塩、鏡像異性体、ジアステレオマー、溶媒和物、または多形体であってもよい。さらに、本明細書に記載される態様または実施形態のいずれかにおいて、本明細書に記載されるULM(または存在する場合、ULM’)は、結合を介して直接、または化学リンカーにより、PTMに結合されてもよい。 In any of the aspects or embodiments described herein, the ULM described herein (or ULM', if present) may be a pharma- ceutically acceptable salt, enantiomer, diastereomer, solvate, or polymorph thereof. Additionally, in any of the aspects or embodiments described herein, the ULM described herein (or ULM', if present) may be attached to the PTM directly via a bond or by a chemical linker.
本開示の特定の態様において、ULM部分は、

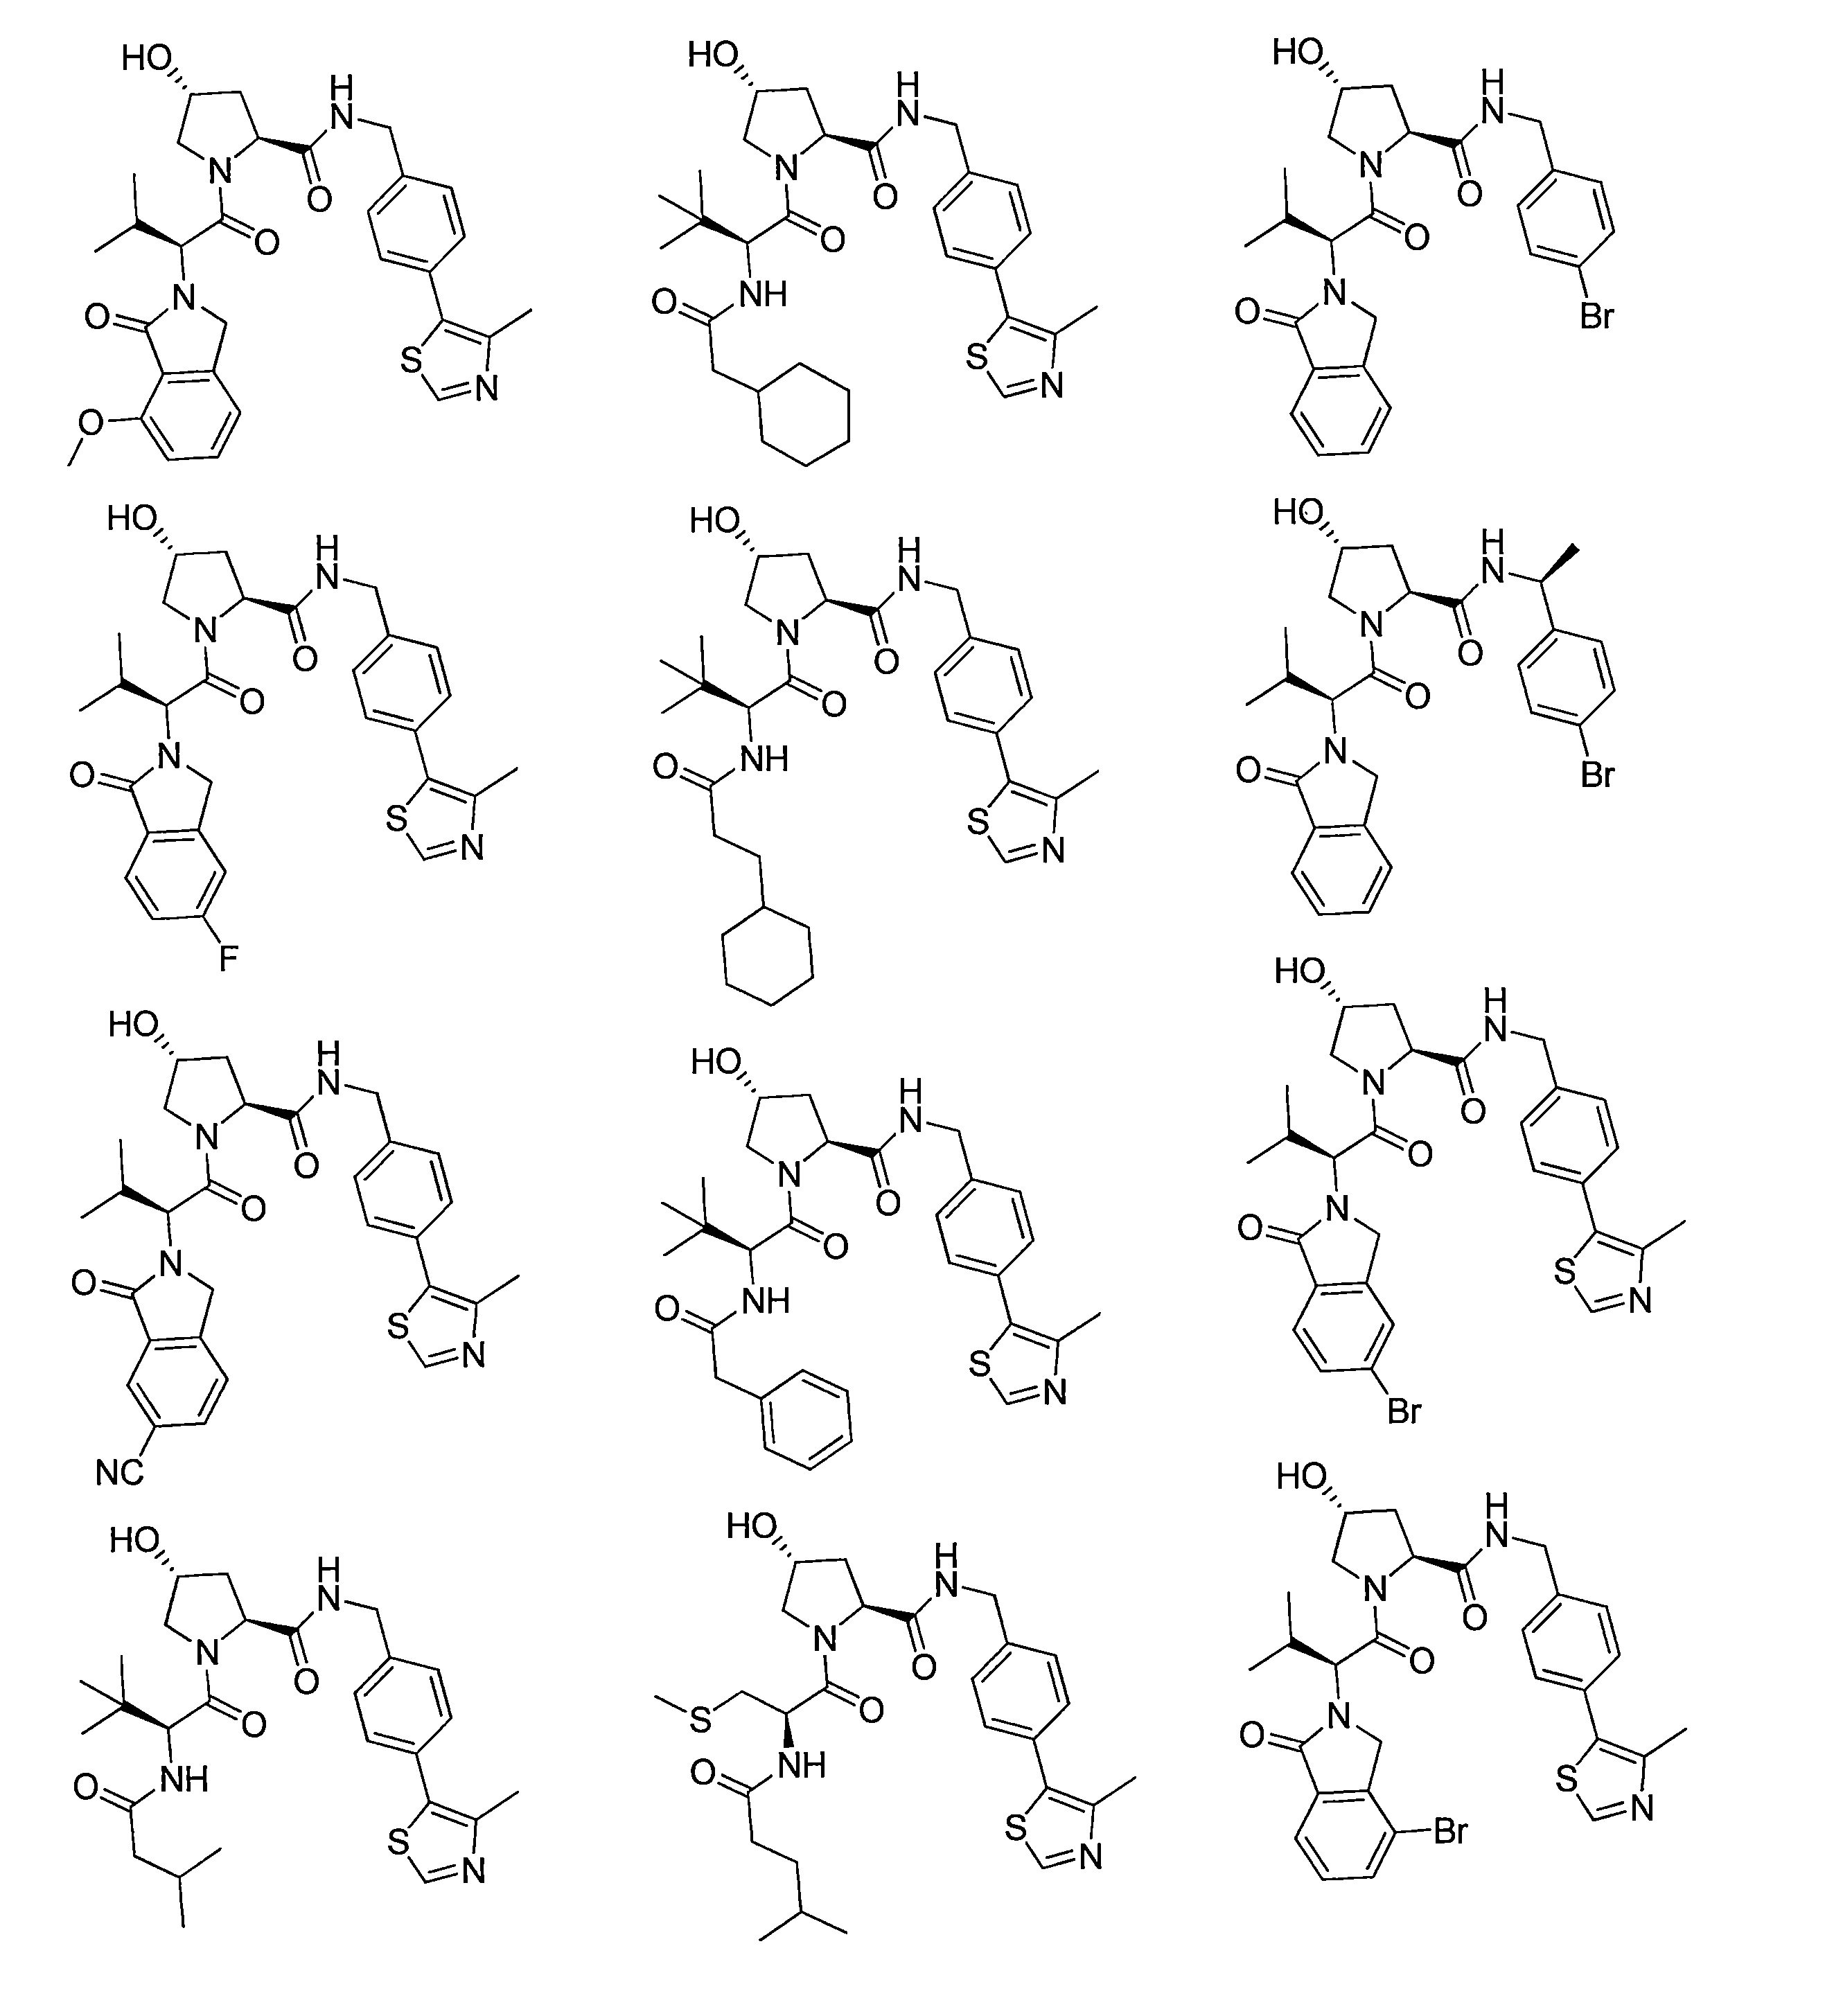
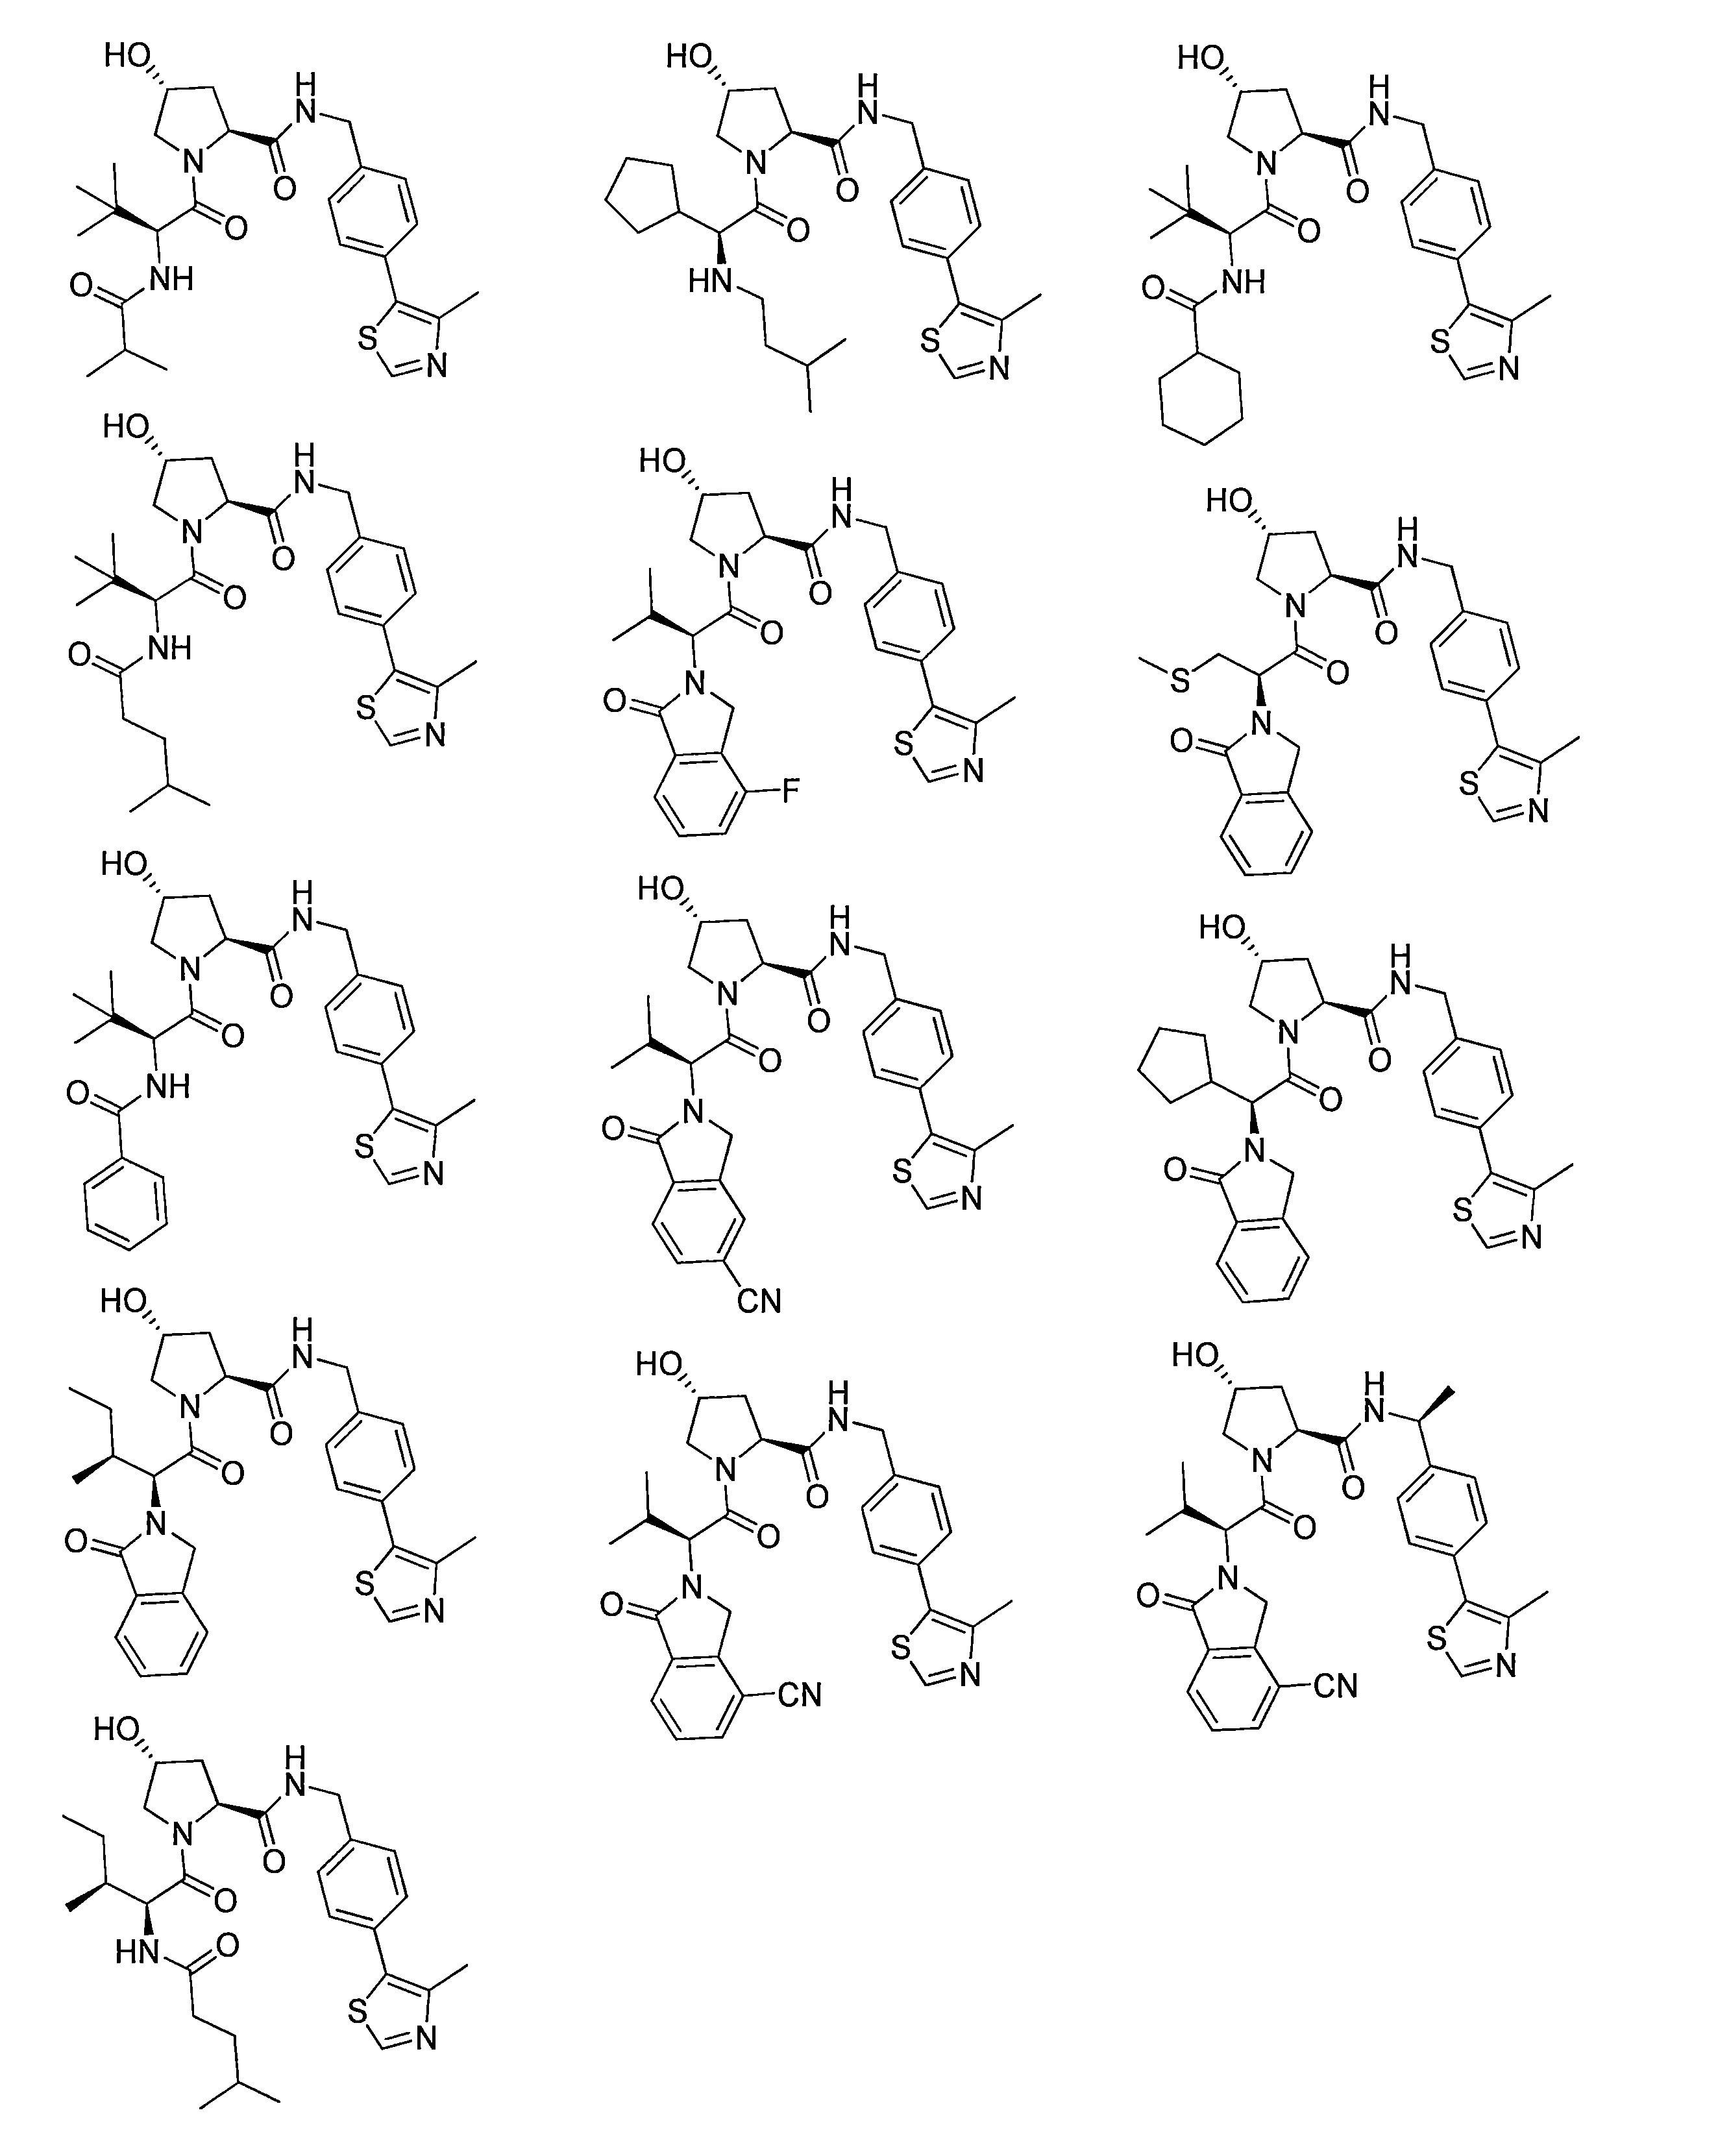


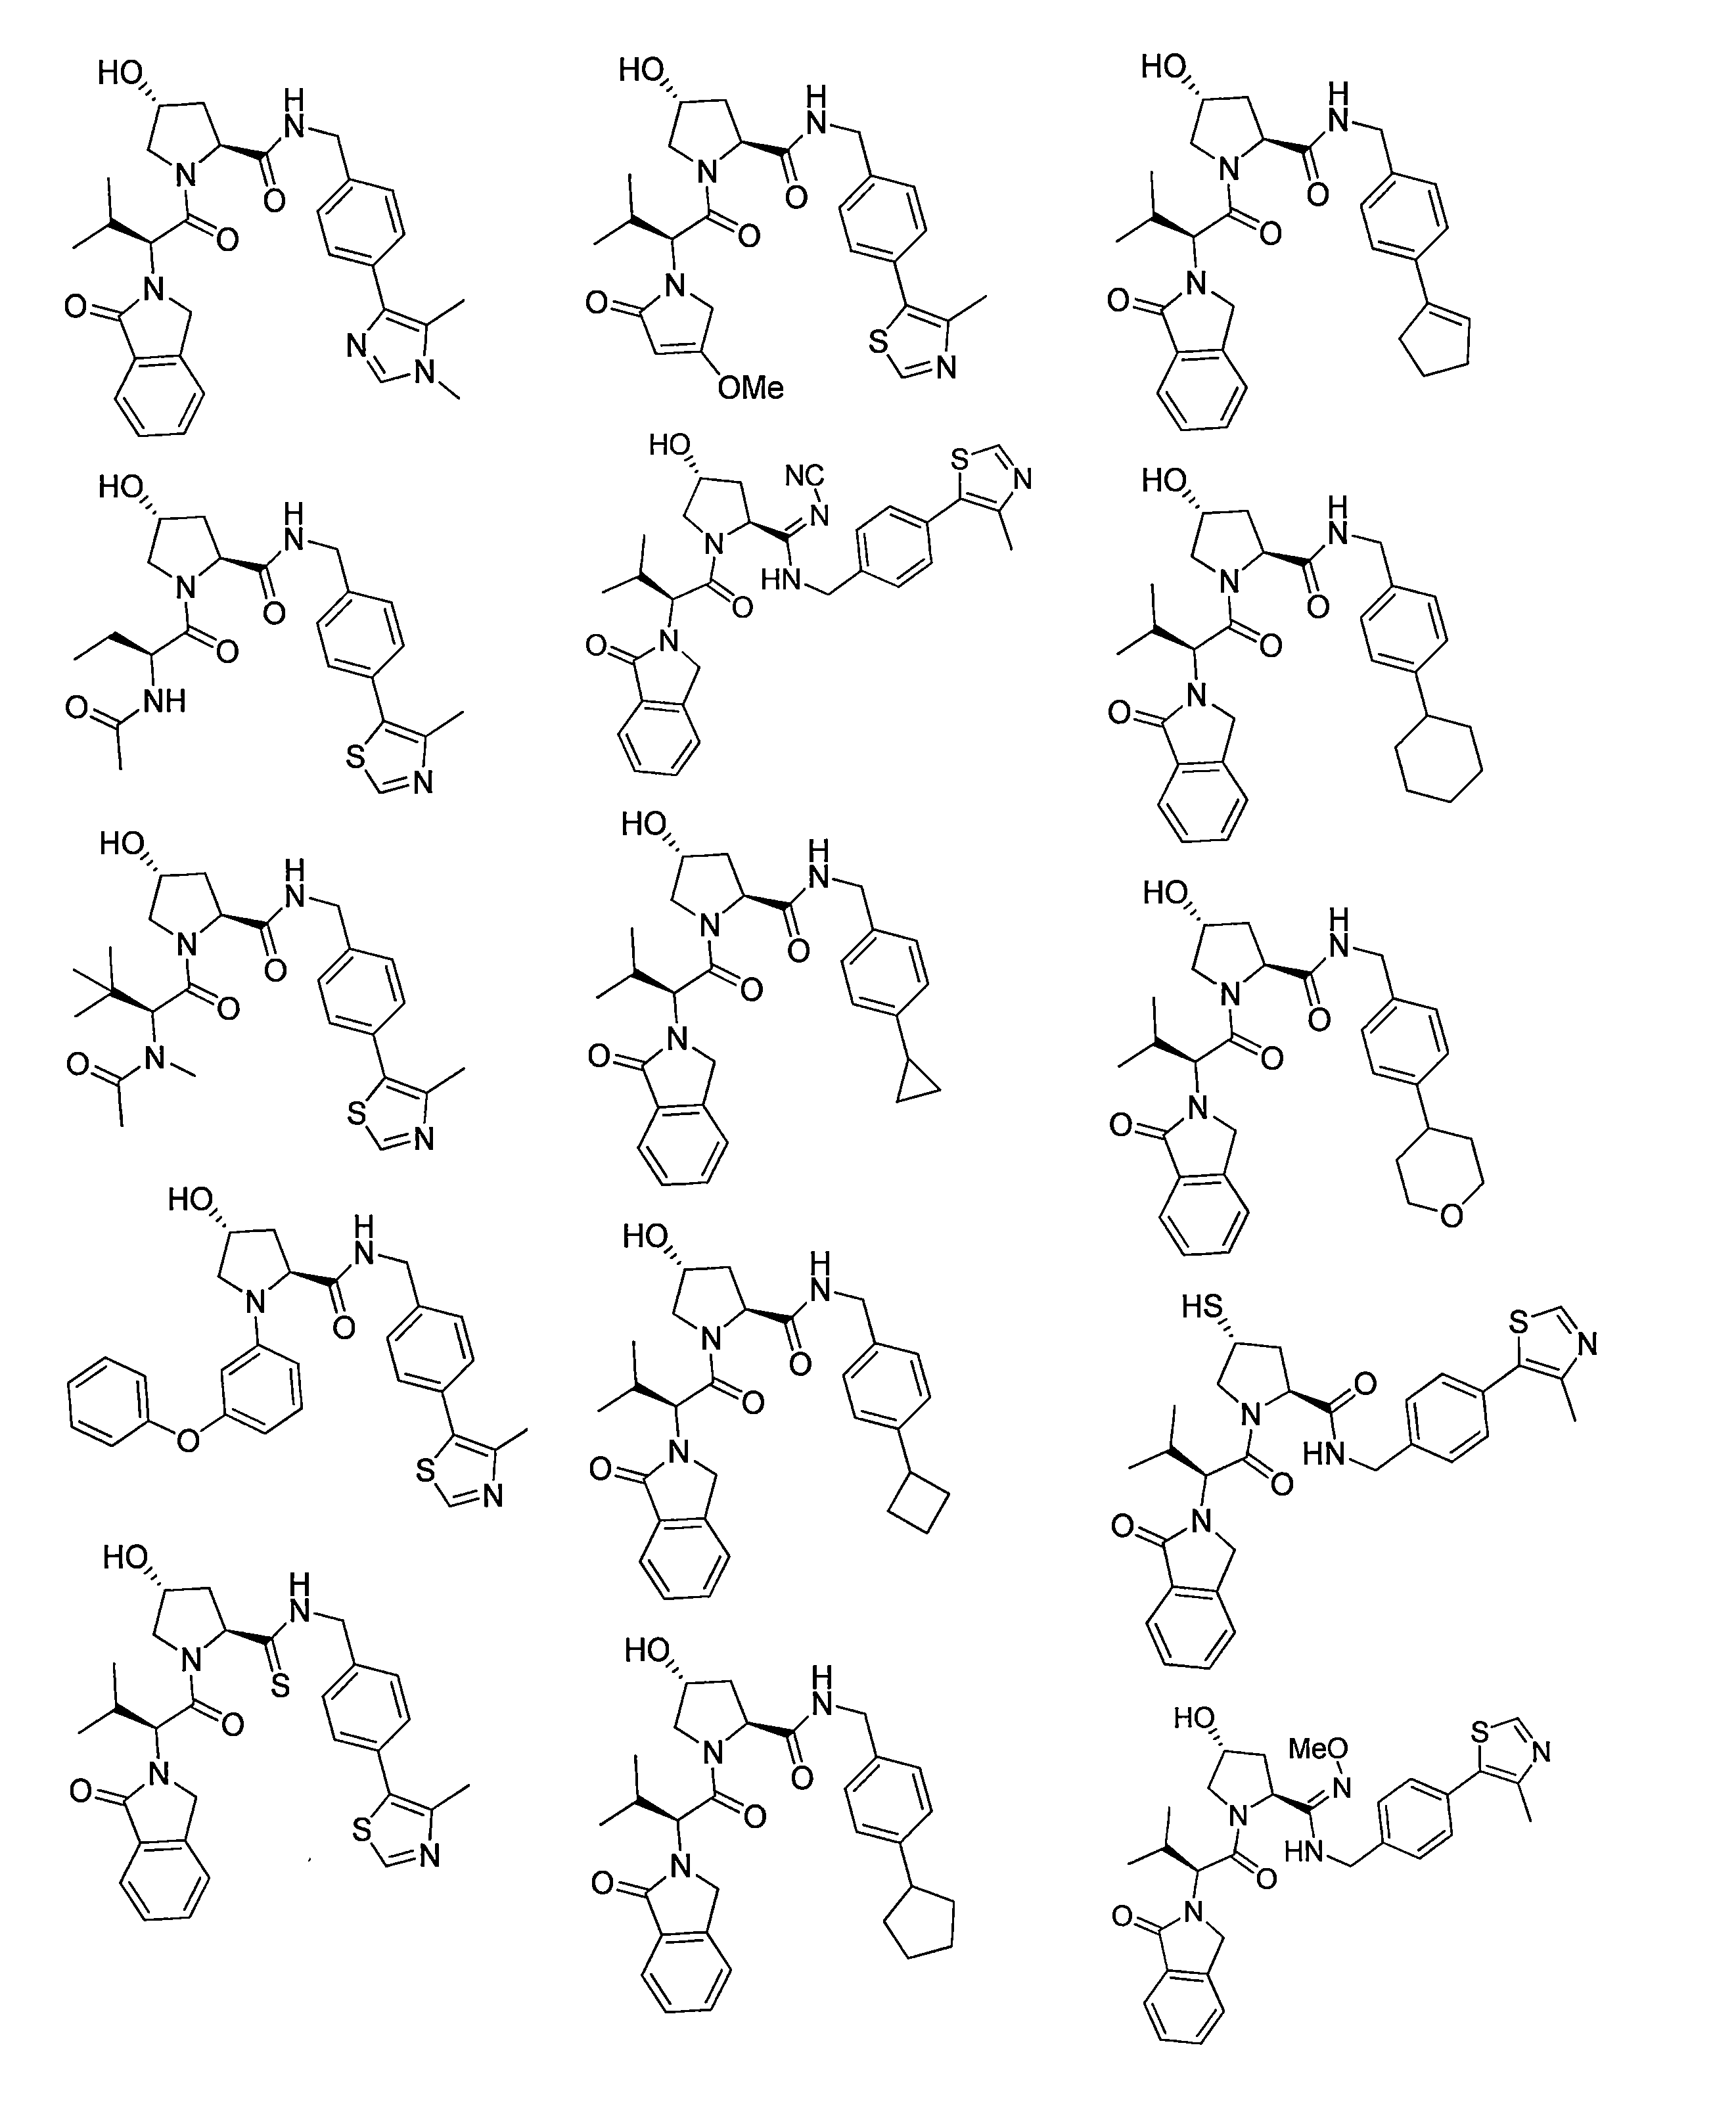


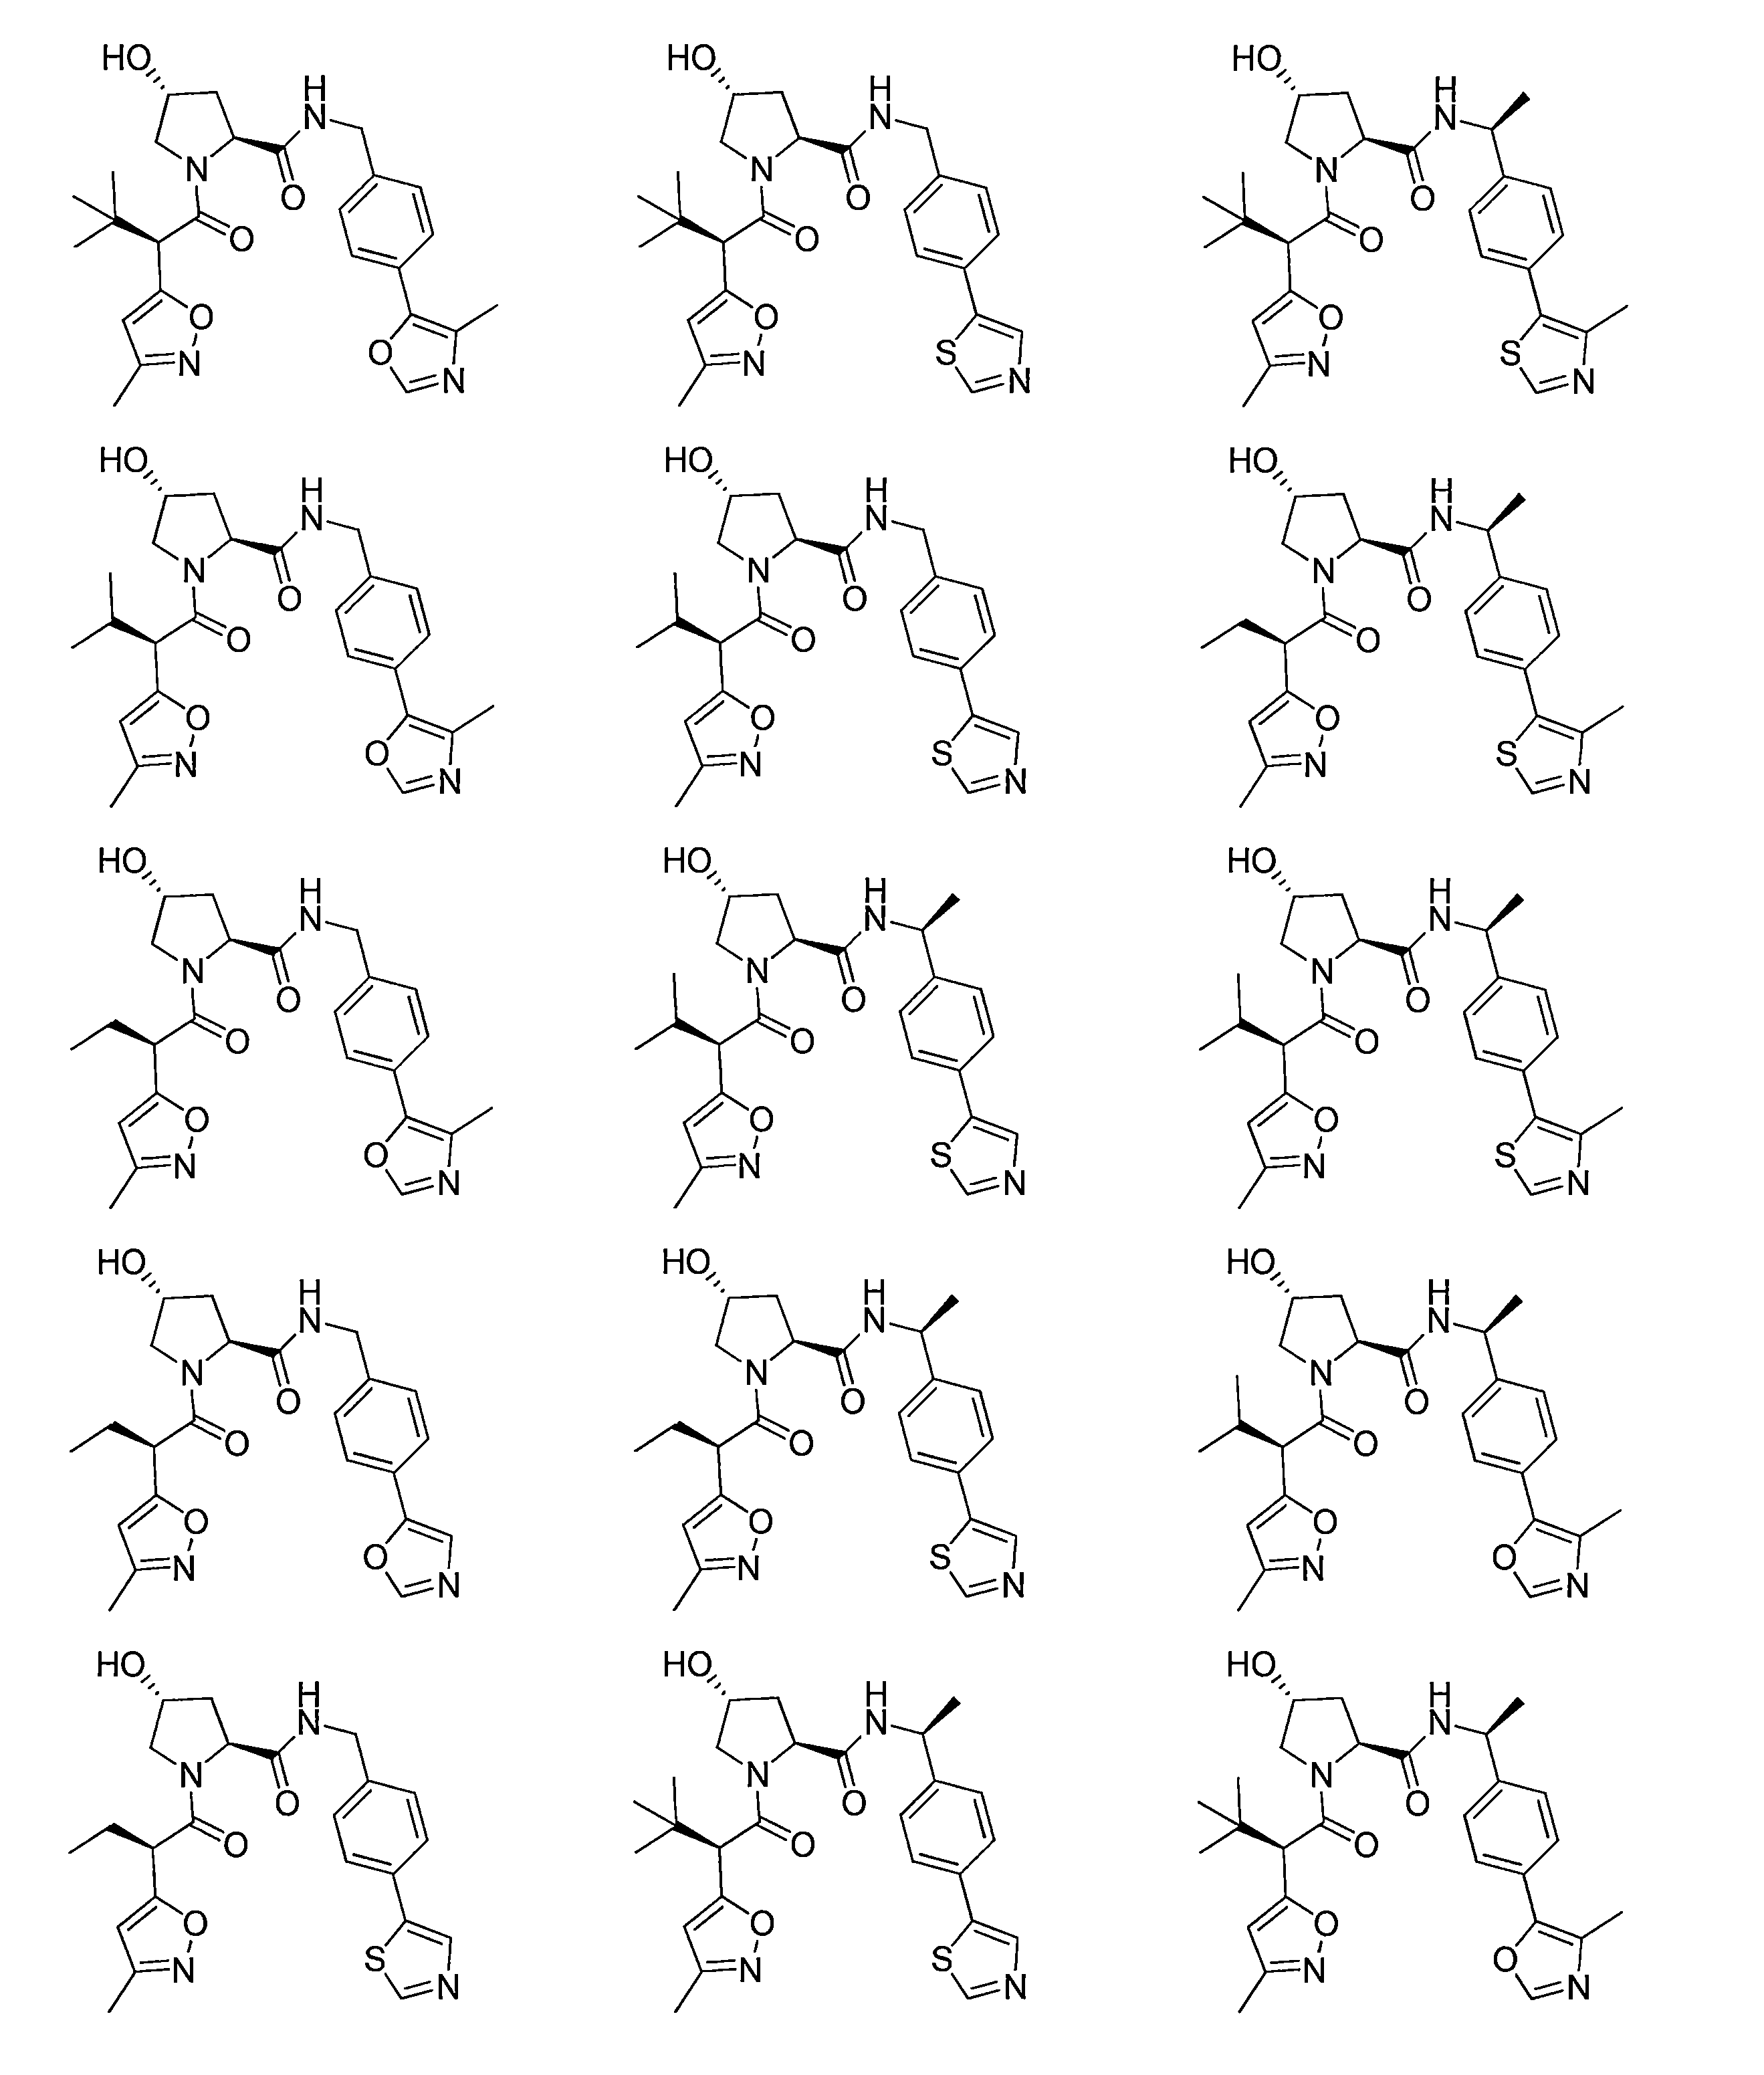
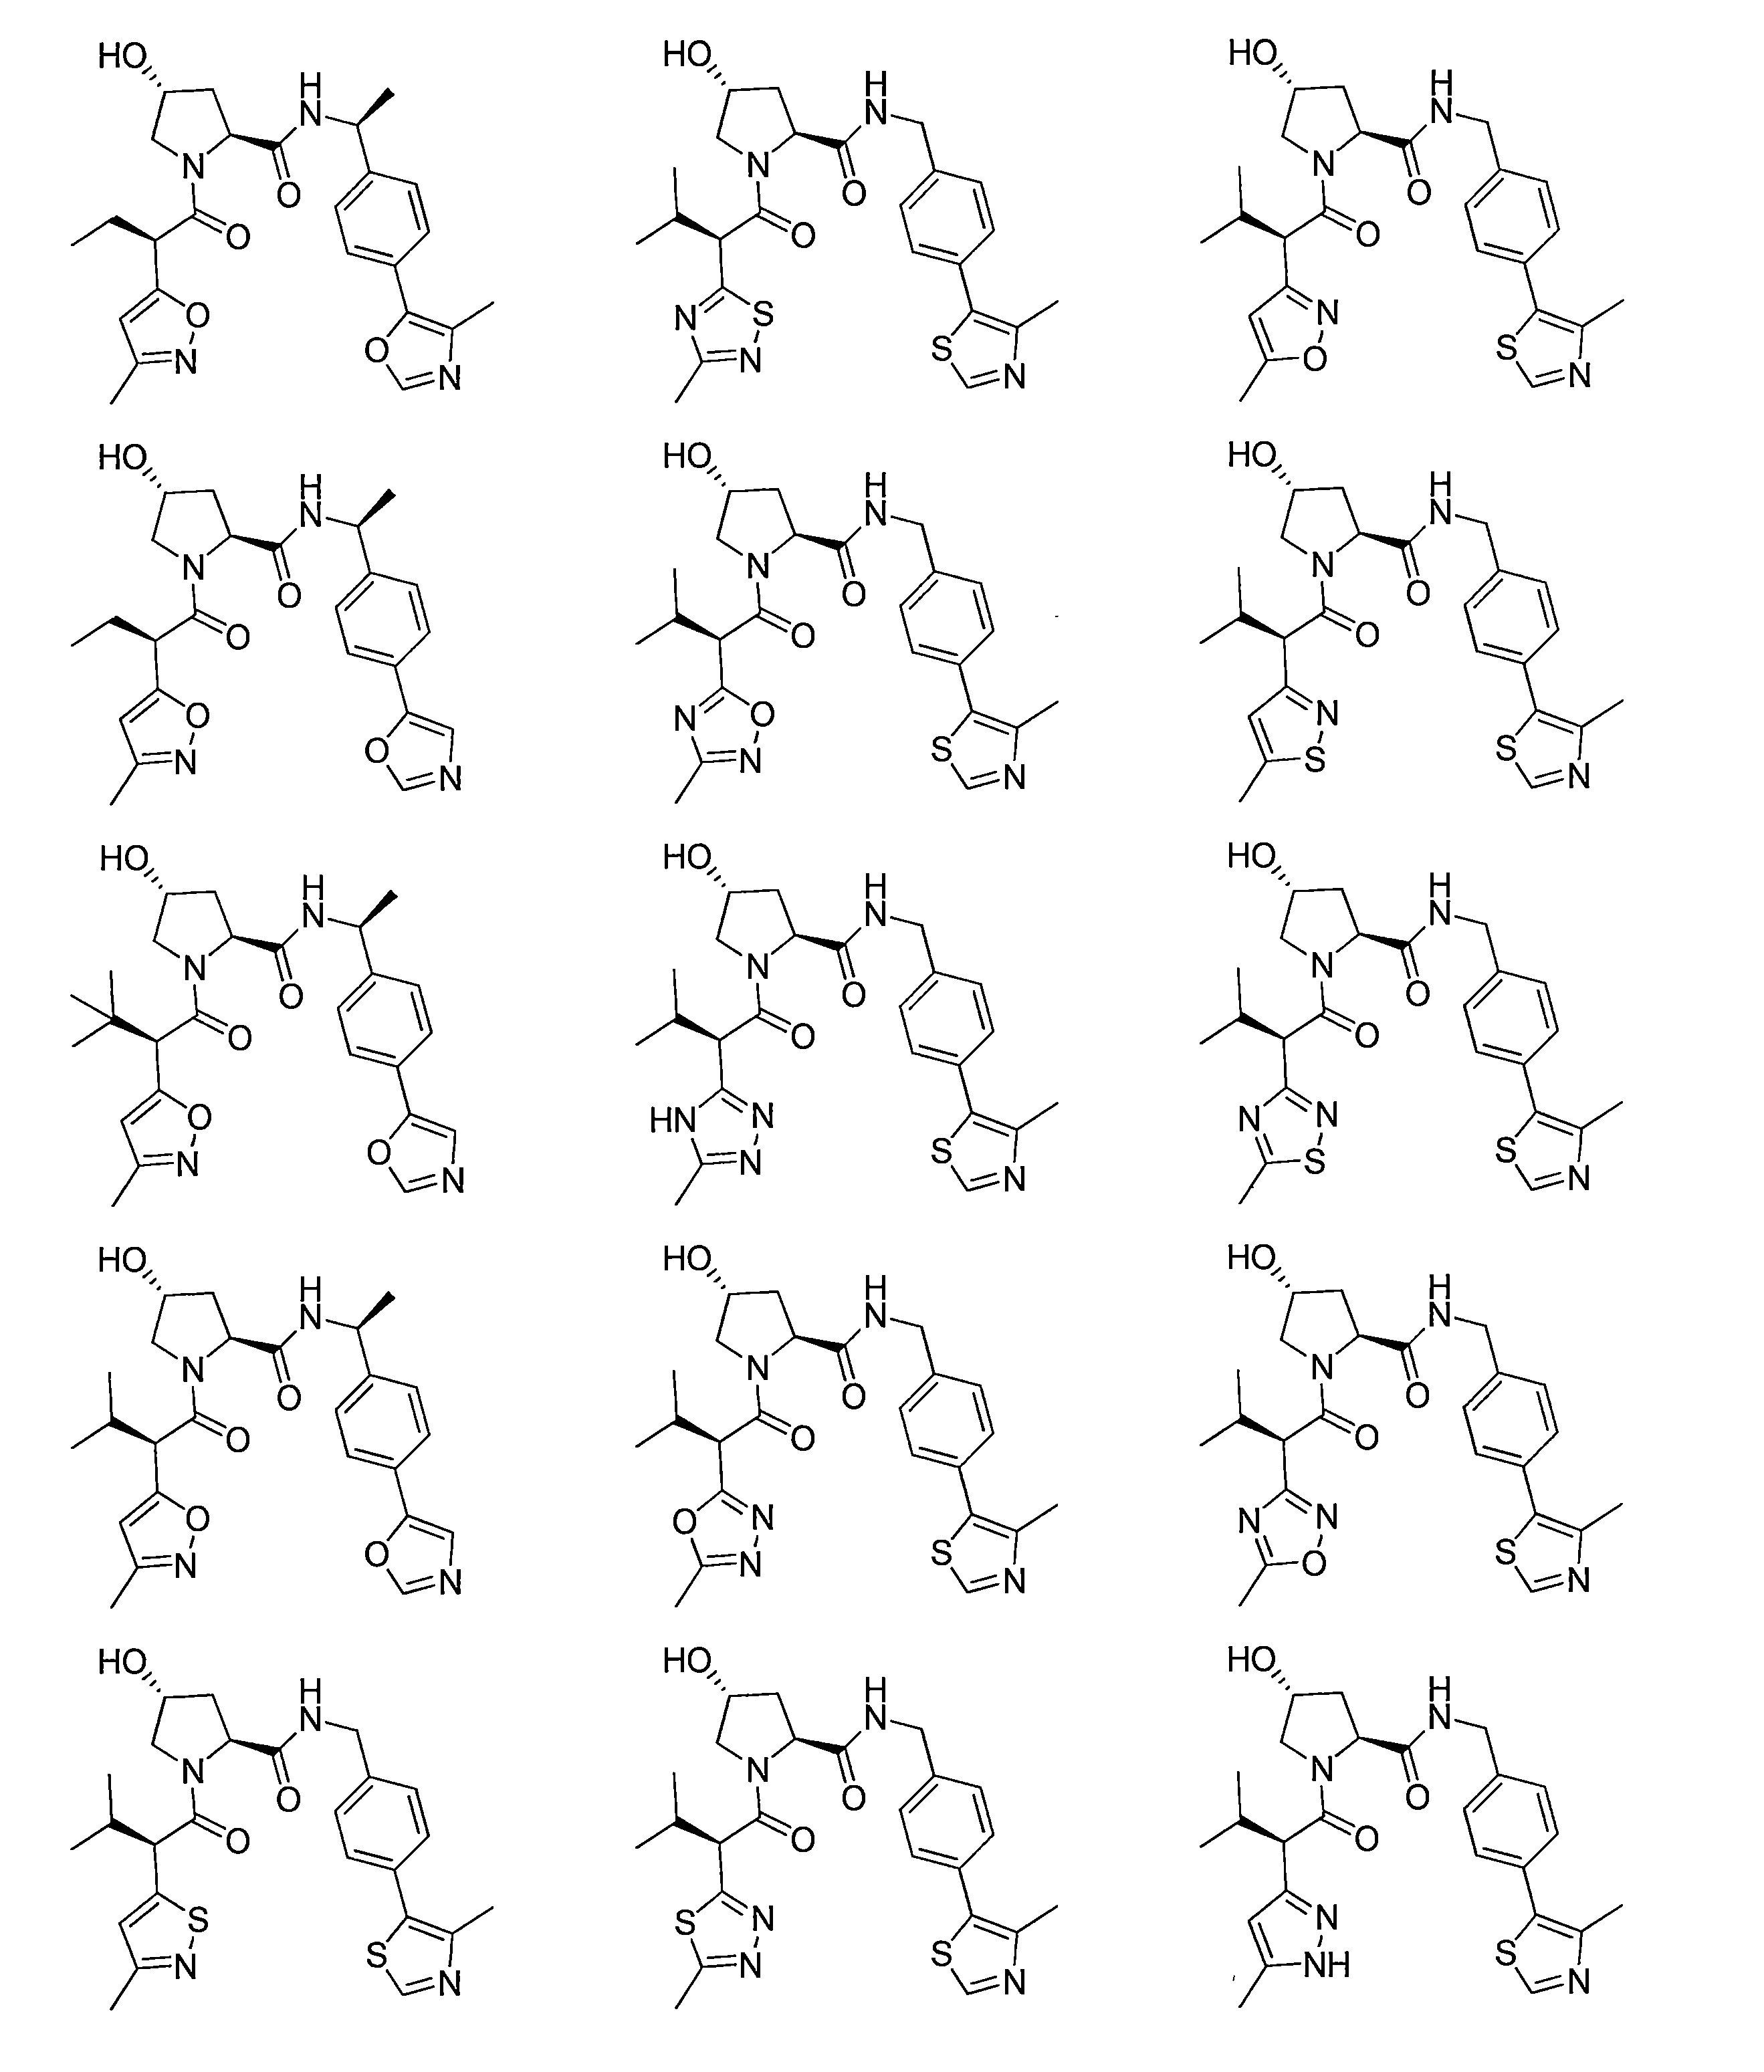

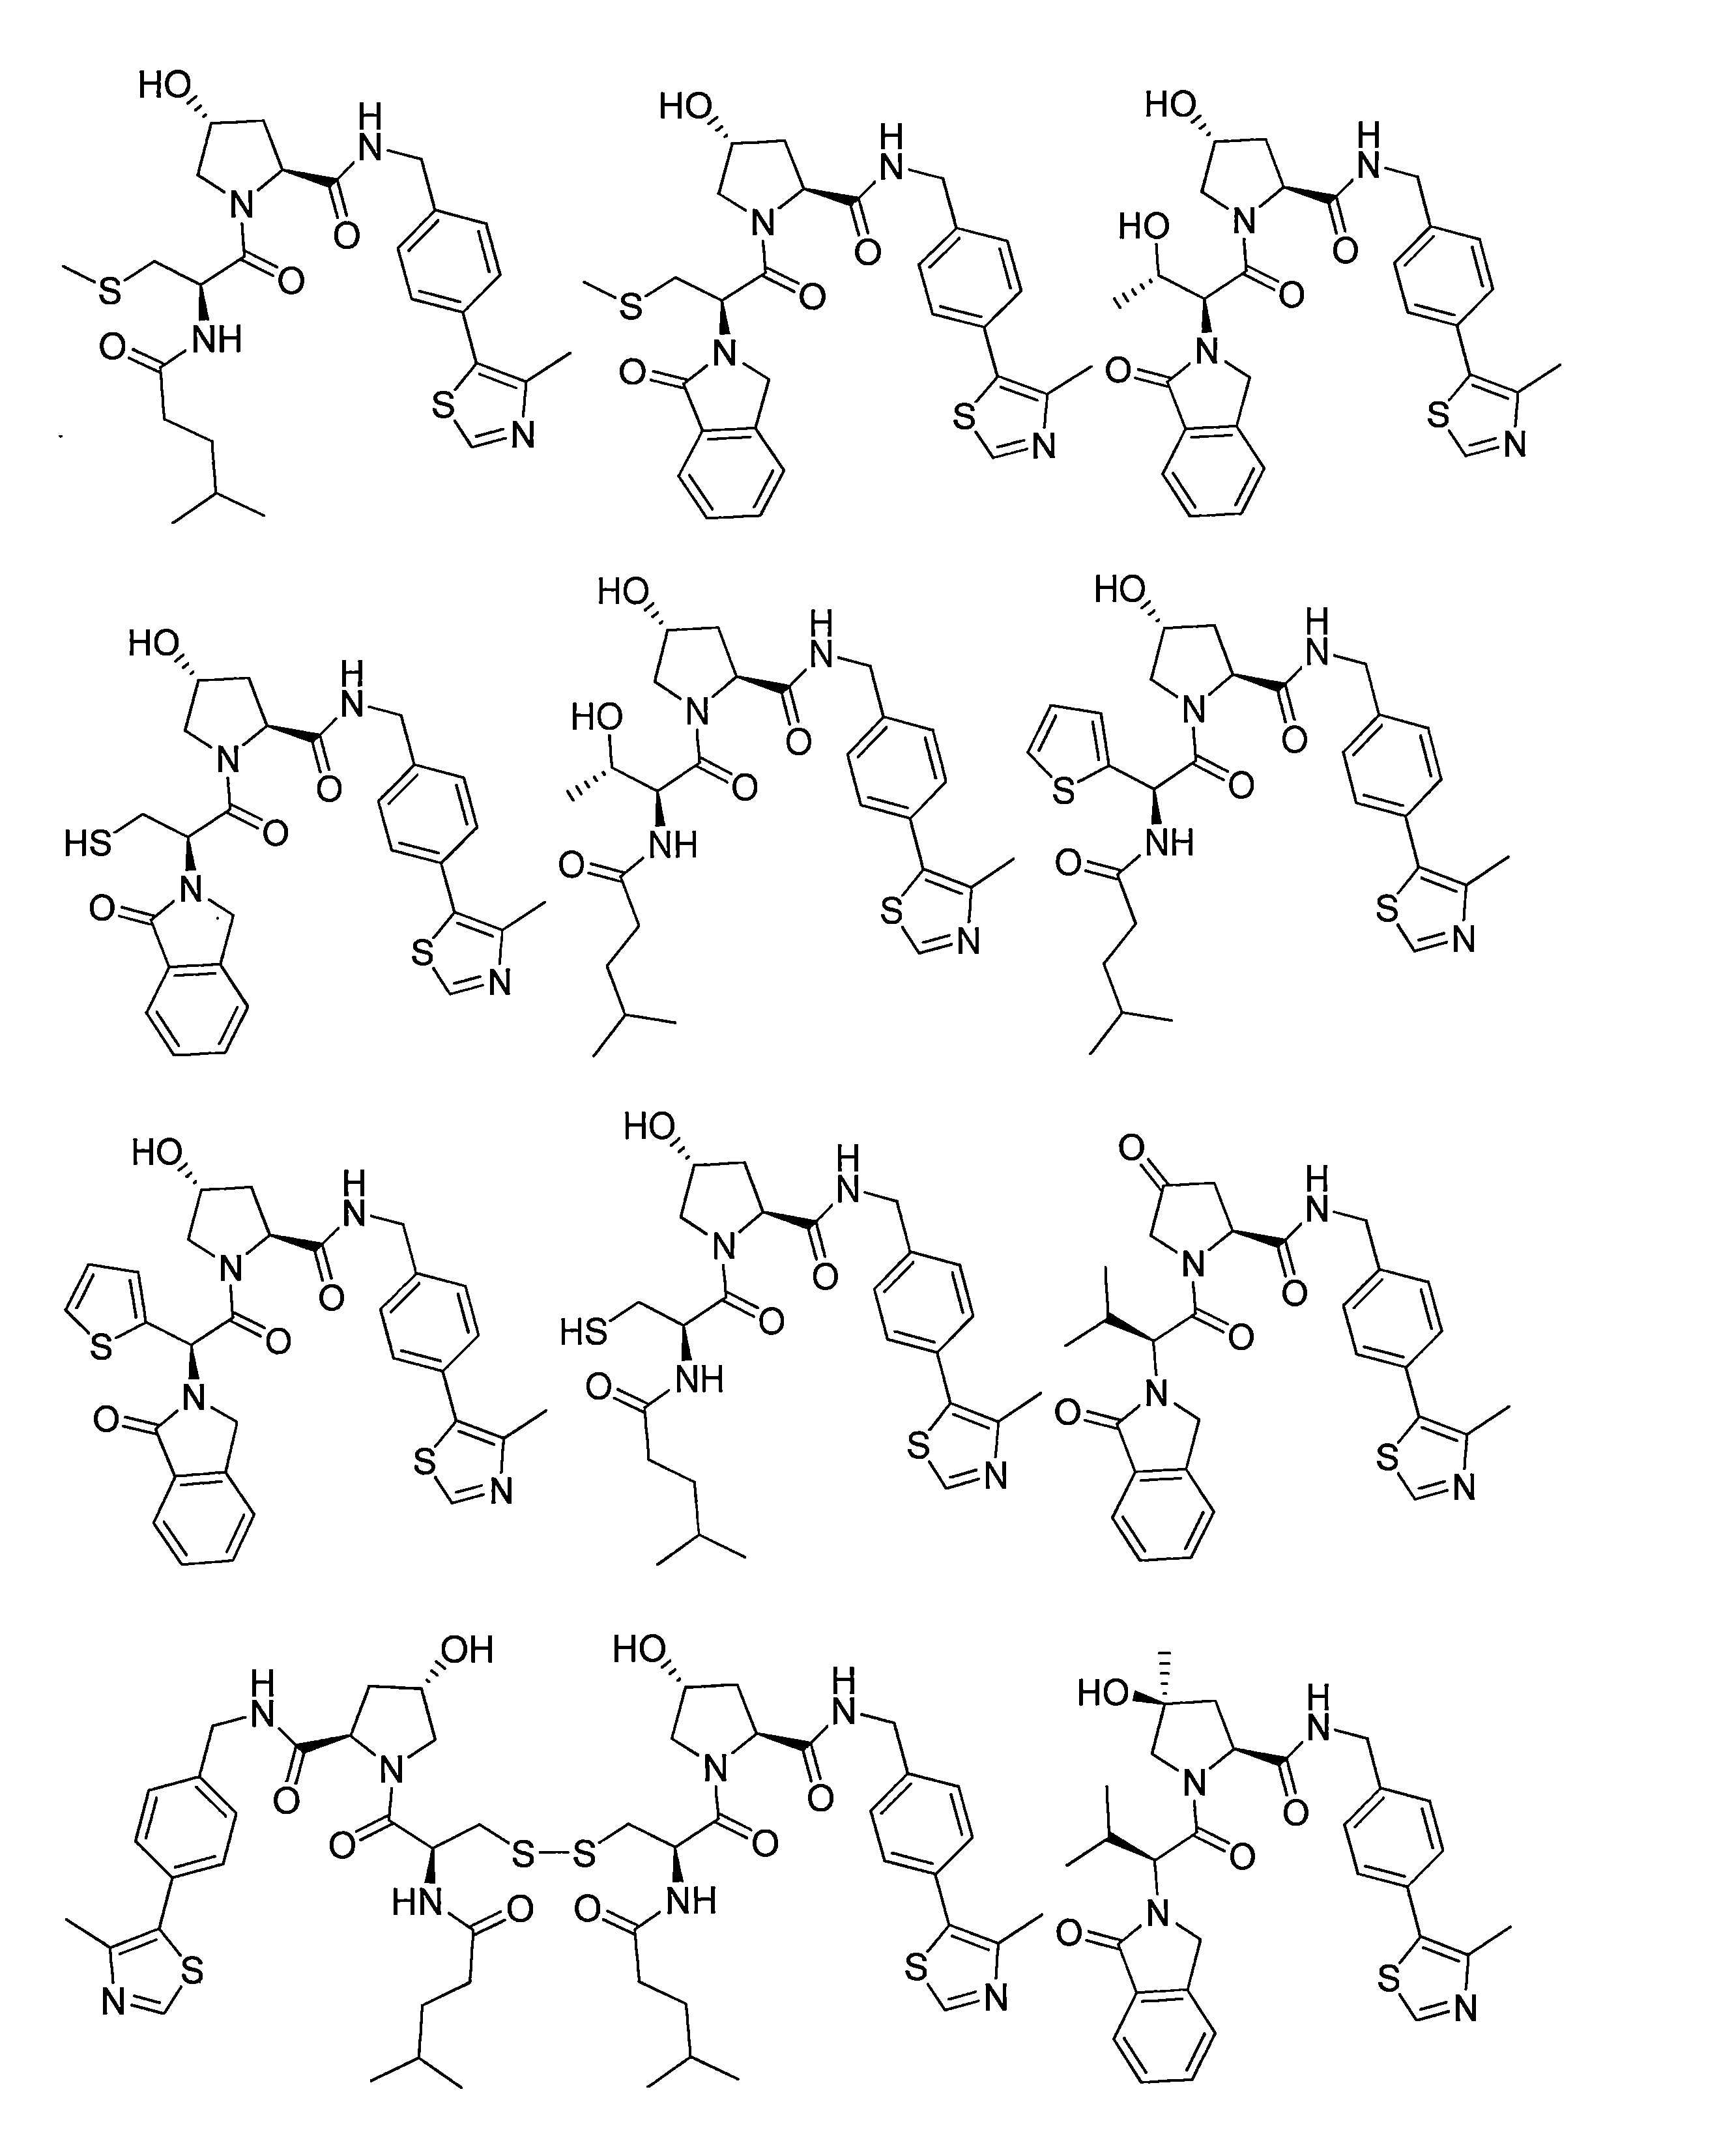


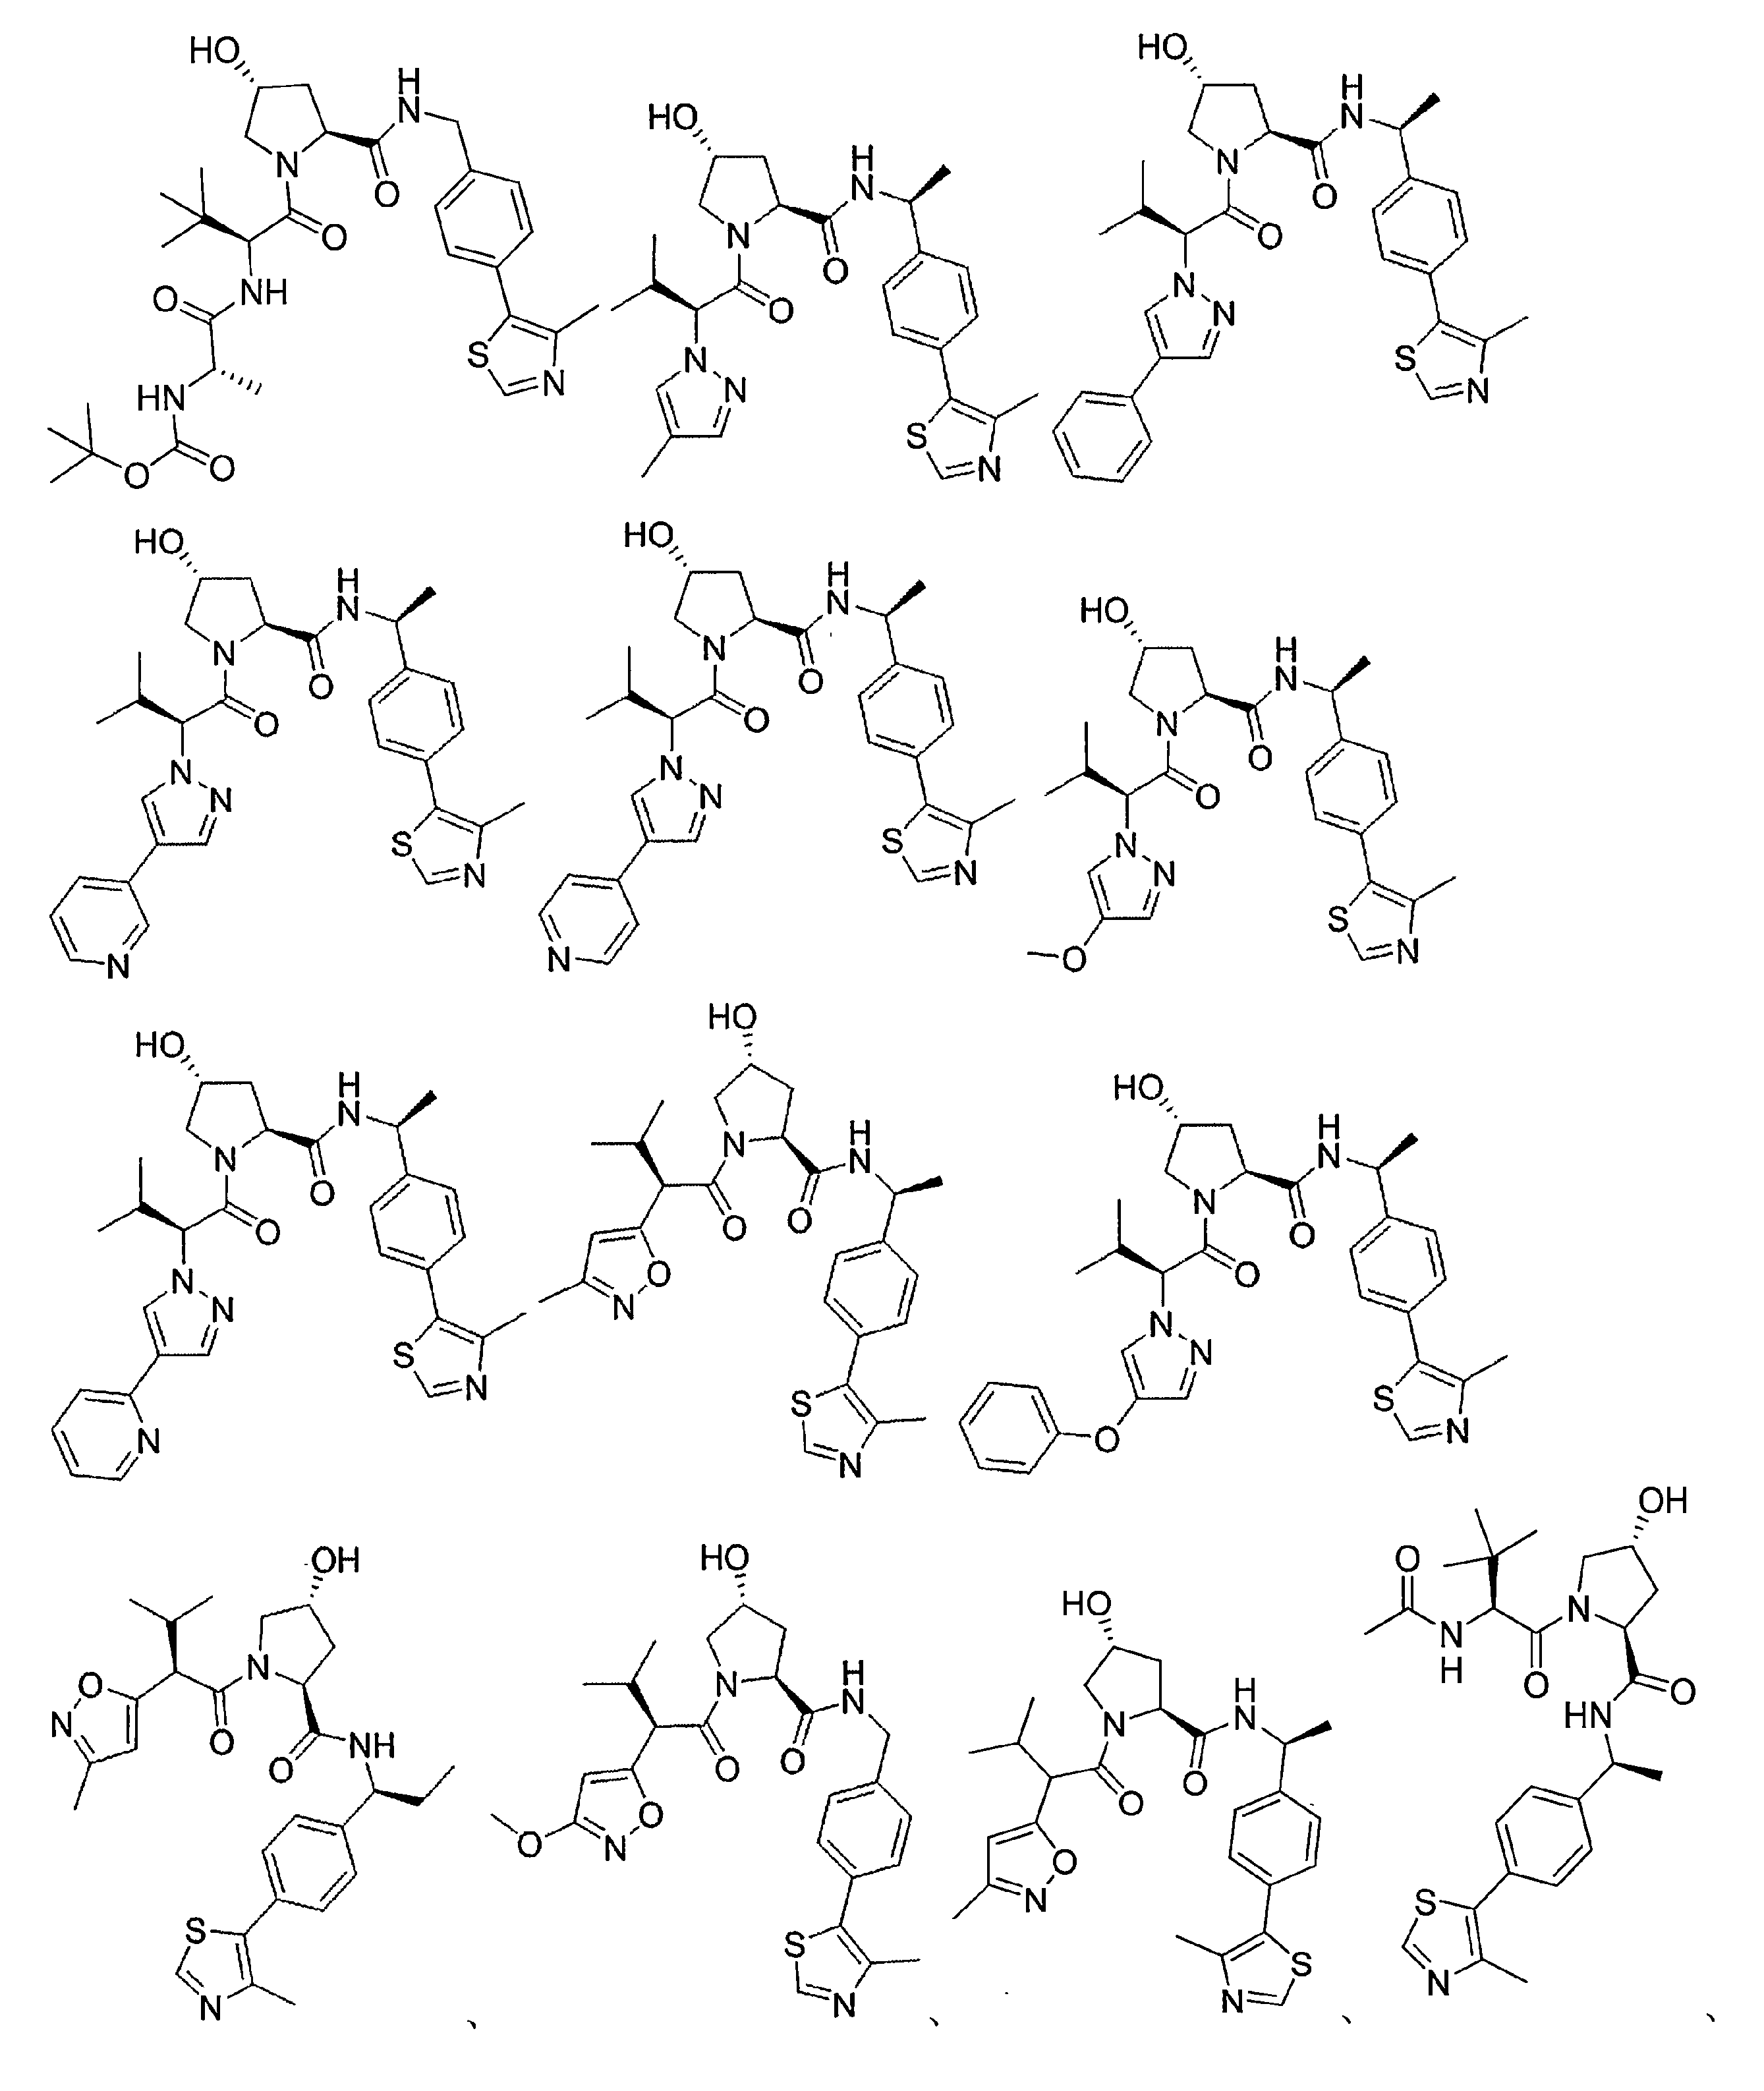

式中、VLMは、本明細書に記載されるリンカーを介して、任意に、任意の適切な官能基、例えば、アミン、エスター、エーテル、アルキル、またはアルコキシを介して、例えば、アリール、ヘテロアリール、フェニル、またはインドール基のフェニルを含む任意の適切な場所で、PTMに結合され得る。
In certain aspects of the present disclosure, the ULM moiety comprises:
wherein the VLM may be attached to the PTM via a linker as described herein, optionally via any suitable functional group, e.g., an amine, ester, ether, alkyl, or alkoxy, at any suitable location, including, e.g., an aryl, heteroaryl, phenyl, or phenyl of an indole group.
例示的リンカー
特定の実施形態では、本明細書に記載される化合物は、PTMをULMに化学結合させるための手段、例えば、化学リンカー(L)を介して1つ以上のULM(例えば、VLMのうちの少なくとも1つ)に化学結合または結合された1つ以上のPTMを含む。ある特定の実施形態では、リンカー基Lは、1つ以上の共有結合した構造単位(例えば、-AL1…(AL)q-または-(AL)q-)を含む基であり、AL1は、PTMに結合した基であり、(AL)qは、ULMに結合した基である。
Exemplary Linkers In certain embodiments, the compounds described herein include one or more PTMs that are chemically bonded or attached to one or more ULMs (e.g., at least one of the VLMs) via a means for chemically linking the PTM to the ULM, e.g., a chemical linker (L). In certain embodiments, the linker group L is a group that includes one or more covalently linked structural units (e.g., -AL1...(AL)q- or -(AL)q-), where AL1 is a group bonded to the PTM and (AL)q is a group bonded to the ULM.
本明細書に記載される任意の態様または実施形態において、リンカー(L)からULM(例えば、VLM、ILM、CLM、またはMLM)への接続または結合は、安定的なL-ULM接続である。例えば、本明細書に記載される任意の態様または実施形態において、リンカー(L)とULMが、ヘテロ原子を介して接続されるとき、後続する任意のヘテロ原子は、存在する場合、例えばアセタール基またはアミナル基など、少なくとも1つの単一炭素原子(例えば、-CH2-)によって分離される。さらなる例として、本明細書に記載される任意の態様または実施形態において、リンカー(L)とULMが、ヘテロ原子を介して接続されるとき、当該ヘテロ原子はエステルの一部ではない。 In any aspect or embodiment described herein, the connection or bond from the linker (L) to the ULM (e.g., VLM, ILM, CLM, or MLM) is a stable L-ULM connection. For example, in any aspect or embodiment described herein, when the linker (L) and the ULM are connected through a heteroatom, any subsequent heteroatom, if present, is separated by at least one single carbon atom (e.g., -CH2-), such as, for example, an acetal or aminal group. As a further example, in any aspect or embodiment described herein, when the linker (L) and the ULM are connected through a heteroatom, the heteroatom is not part of an ester.
本明細書に記載される任意の態様または実施形態では、リンカー基Lは、式-(AL)q-によって表される結合または化学リンカー基であり、式中、Aが、化学部分であり、qが、1~100の整数であり、Lが、PTMおよびULMに共有結合され、PTMのタンパク質標的への、およびULMのE3ユビキチンへの十分な結合を提供して、標的タンパク質ユビキチン化をもたらす。 In any aspect or embodiment described herein, the linker group L is a bond or chemical linker group represented by the formula -(AL)q-, where A is a chemical moiety and q is an integer from 1 to 100, and L is covalently attached to the PTM and ULM and provides sufficient binding of the PTM to a protein target and the ULM to E3 ubiquitin to effect target protein ubiquitination.
本明細書に記載される任意の態様または実施形態では、リンカー基Lは、-(AL)q-であり、式中、
(AL)qは、ULM(例えばVLM)、PTM部分、またはそれらの組み合わせのうちの少なくとも1つに連結された基であり、
リンカーのqは、1以上の整数であり、
各ALは、独立して、結合、CRL1RL2、O、S、SO、SO2、NRL3、SO2NRL3、SONRL3、CONRL3、NRL3CONRL4、NRL3SO2NRL4、CO、CRL1=CRL2、C≡C、SiRL1RL2、P(O)RL1、P(O)ORL1、NRL3C(=NCN)NRL4、NRL3C(=NCN)、NRL3C(=CNO2)NRL4、0~6個のRL1および/またはRL2基で任意に置換されたC3-11シクロアルキル、0~9個のRL1および/またはRL2基で任意に置換されたC5-13スピロシクロアルキル、0~6個のRL1および/またはRL2基で任意に置換されたC3-11ヘテロシクリル、0~8個のRL1および/またはRL2基で任意に置換されたC5-13スピロヘテロシクリル、0~6個のRL1および/またはRL2基で任意に置換されたアリール、0~6個のRL1および/またはRL2基で任意に置換されたヘテロアリールからなる群から選択され、式中、RL1またはRL2は、各々独立して、任意に他の基に連結されて、シクロアルキルおよび/またはヘテロシクリル部分を形成し、任意に0~4個のRL5基で置換され、
RL1、RL2、RL3、RL4およびRL5は、各々独立して、H、ハロ、C1-8アルキル、OC1-8アルキル、SC1-8アルキル、NHC1-8アルキル、N(C1-8アルキル)2、C3-11シクロアルキル、アリール、ヘテロアリール、C3-11ヘテロシクリル、OC1-8シクロアルキル、SC1-8シクロアルキル、NHC1-8シクロアルキル、N(C1-8シクロアルキル)2、N(C1-8シクロアルキル)(C1-8アルキル)、OH、NH2、SH、SO2C1-8アルキル、P(O)(OC1-8アルキル)(C1-8アルキル)、P(O)(OC1-8アルキル)2、CC-C1-8アルキル、CCH、CH=CH(C1-8アルキル)、C(C1-8アルキル)=CH(C1-8アルキル)、C(C1-8アルキル)=C(C1-8アルキル)2、Si(OH)3、Si(C1-8アルキル)3、Si(OH)(C1-8アルキル)2、COC1-8アルキル、CO2H、ハロゲン、CN、CF3、CHF2、CH2F、NO2、SF5、SO2NHC1-8アルキル、SO2N(C1-8アルキル)2、SONHC1-8アルキル、SON(C1-8アルキル)2、CONHC1-8アルキル、CON(C1-8アルキル)2、N(C1-8アルキル)CONH(C1-8アルキル)、N(C1-8アルキル)CON(C1-8アルキル)2、NHCONH(C1-8アルキル)、NHCON(C1-8アルキル)2、NHCONH2、N(C1-8アルキル)SO2NH(C1-8アルキル)、N(C1-8アルキル)SO2N(C1-8アルキル)2、NH SO2NH(C1-8アルキル)、NH SO2N(C1-8アルキル)2、NH SO2NH2である。
In any aspect or embodiment described herein, the linker group L is -(AL)q-, wherein:
(AL)q is a group linked to at least one of a ULM (e.g., VLM), a PTM moiety, or a combination thereof;
The q of the linker is an integer of 1 or more,
each AL is independently a bond, CRL1RL2, O, S, SO, SO2, NRL3, SON2NRL3, SONRL3, CONRL3, NRL3CONRL4, NRL3SON2NRL4, CO, CRL1=CRL2, C≡C, SiRL1RL2, P(O)RL1, P(O)ORL1, NRL3C(=NCN)NRL4, NRL3C(=NCN), NRL3C(=CNO2)NRL4, C3-11 cycloalkyl optionally substituted with 0-6 RL1 and/or RL2 groups, C5-13 spirocyclo optionally substituted with 0-9 RL1 and/or RL2 groups, alkyl, C3-11 heterocyclyl optionally substituted with 0-6 R and/or R groups, C5-13 spiroheterocyclyl optionally substituted with 0-8 R and/or R groups, aryl optionally substituted with 0-6 R and/or R groups, heteroaryl optionally substituted with 0-6 R and/or R groups, wherein R or R are each independently optionally linked to other groups to form a cycloalkyl and/or heterocyclyl moiety and are optionally substituted with 0-4 R groups;
RL1, RL2, RL3, RL4 and RL5 are each independently H, halo, C1-8 alkyl, OC1-8 alkyl, SC1-8 alkyl, NHC1-8 alkyl, N(C1-8 alkyl)2, C3-11 cycloalkyl, aryl, heteroaryl, C3-11 heterocyclyl, OC1-8 cycloalkyl, SC1-8 cycloalkyl, NHC1-8 cycloalkyl, N(C1-8 cycloalkyl)2, N(C1-8 cycloalkyl)(C1-8 alkyl), OH, NH2, SH, SO2C1-8 alkyl, P(O)(OC1-8 alkyl)(C1-8 alkyl), P(O)(OC1-8 alkyl)2, CC-C1-8 alkyl, CCH, CH=CH(C1-8 alkyl), C(C1-8 alkyl)=CH(C1-8 alkyl), C(C1-8 alkyl), alkyl) = C(C1-8 alkyl)2, Si(OH)3, Si(C1-8 alkyl)3, Si(OH)(C1-8 alkyl)2, COC1-8 alkyl, CO2H, halogen, CN, CF3, CHF2, CH2F, NO2, SF5, SON2NHC1-8 alkyl, SON2N(C1-8 alkyl)2, SONHC1-8 alkyl, SON(C1-8 alkyl)2, CONHC1-8 alkyl, CON(C1-8 alkyl)2, N(C1-8 alkyl)CONH(C1-8 alkyl), N(C1-8 alkyl)CON(C1-8 alkyl)2, NHCONH(C1-8 alkyl), NHCON(C1-8 alkyl)2, NHCONH2, N(C1-8 alkyl)SON2NH(C1-8 alkyl), N(C1-8 alkyl)SON2N(C1-8 alkyl)2, NH SO2NH(C1-8 alkyl), NHSO2N(C1-8 alkyl)2, NHSO2NH2.
特定の実施形態では、リンカーのqは0以上の整数である。特定の実施形態では、qは1以上の整数である。 In certain embodiments, q in the linker is an integer greater than or equal to 0. In certain embodiments, q is an integer greater than or equal to 1.
例えば、リンカーのqが2より大きい、ある特定の実施形態では、(AL)qは、AL1および(AL)qである基であり、単位ALは、PTMをULMに結合させる。 For example, in certain embodiments where the linker q is greater than 2, (AL)q is a group that is AL1 and (AL)q, and the unit AL connects the PTM to the ULM.
例えば、リンカーのqが2である、ある特定の実施形態では、(AL)qは、AL1に、およびULMまたはPTMに連結している基である。 For example, in certain embodiments where the linker q is 2, (AL)q is a group that links to AL1 and to the ULM or PTM.
例えば、リンカーのqが1である、ある特定の実施形態では、リンカー基Lの構造は、-AL1-であり、AL1は、ULM部分およびPTM部分に連結している基である。 For example, in certain embodiments where the linker q is 1, the structure of the linker group L is -AL1-, where AL1 is a group that connects the ULM and PTM moieties.
ある特定の実施形態では、リンカー(L)の単位ALは、以下からなる群から選択される一般構造によって表される基を含み、
-NR(CH2)n-(低級アルキル)-、-NR(CH2)n-(低級アルコキシル)-、-NR(CH2)n-(低級アルコキシル)-OCH2-、-NR(CH2)n-(低級アルコキシル)-(低級アルキル)-OCH2-、-NR(CH2)n-(シクロアルキル)-(低級アルキル)-OCH2-、-NR(CH2)n-(ヘテロシクロアルキル)-、-NR(CH2CH2O)n-(低級アルキル)-O-CH2-、-NR(CH2CH2O)n-(ヘテロシクロアルキル)-O-CH2-、-NR(CH2CH2O)n-アリール-O-CH2-、-NR(CH2CH2O)n-(ヘテロアリール)-O-CH2-、-NR(CH2CH2O)n-(シクロアルキル)-O-(ヘテロアリール)-O-CH2-、-NR(CH2CH2O)n-(シクロアルキル)-O-アリール-O-CH2-、-NR(CH2CH2O)n-(低級アルキル)-NH-アリール-O-CH2-、-NR(CH2CH2O)n-(低級アルキル)-O-アリール-CH2、-NR(CH2CH2O)n-シクロアルキル-O-アリール-、-NR(CH2CH2O)n-シクロアルキル-O-(ヘテロアリール)l-、-NR(CH2CH2)n-(シクロアルキル)-O-(ヘテロシクリル)-CH2、-NR(CH2CH2)n-(ヘテロシクリル)-(ヘテロシクリル)-CH2、-N(R1R2)-(ヘテロシクリル)-CH2、式中、
リンカーのnは、0~10であってもよく、
リンカーのRは、H、低アルキルであってもよく、
リンカーのR1およびR2は、N結合を有する環を形成してもよい。
In certain embodiments, the unit AL of the linker (L) comprises a group represented by a general structure selected from the group consisting of:
—NR(CH2)n-(lower alkyl)-, —NR(CH2)n-(lower alkoxyl)-, —NR(CH2)n-(lower alkoxyl)-OCH2-, —NR(CH2)n-(lower alkoxyl)-(lower alkyl)-OCH2-, —NR(CH2)n-(cycloalkyl)-(lower alkyl)-OCH2-, —NR(CH2)n-(heterocycloalkyl)-, —NR(CH2CH2O)n-(lower alkyl)-O-CH2-, —NR(CH2CH2O)n-(heterocycloalkyl)-O-CH2-, —NR(CH2CH2O)n-aryl-O-CH2-, —NR(CH2CH2O)n-(heteroaryl)-O-CH2-, —NR(CH2CH2O)n-(cyclo alkyl)-O-(heteroaryl)-O-CH2-, -NR(CH2CH2O)n-(cycloalkyl)-O-aryl-O-CH2-, -NR(CH2CH2O)n-(lower alkyl)-NH-aryl-O-CH2-, -NR(CH2CH2O)n-(lower alkyl)-O-aryl-CH2, -NR(CH2CH2O)n-cycloalkyl-O-aryl-, -NR(CH2CH2O)n-cycloalkyl-O-(heteroaryl)l-, -NR(CH2CH2)n-(cycloalkyl)-O-(heterocyclyl)-CH2, -NR(CH2CH2)n-(heterocyclyl)-(heterocyclyl)-CH2, -N(RR2)-(heterocyclyl)-CH2,
The linker's n can be 0 to 10;
R of the linker can be H, lower alkyl;
The linkers R1 and R2 may form a ring with the N-bond.
ある特定の実施形態では、リンカー(L)の単位ALは、以下からなる群から選択される一般構造によって表される基を含み、
-N(R)-(CH2)m-O(CH2)n-O(CH2)o-O(CH2)p-O(CH2)q-O(CH2)r-OCH2-、
-O-(CH2)m-O(CH2)n-O(CH2)o-O(CH2)p-O(CH2)q-O(CH2)r-OCH2-、
-O-(CH2)m-O(CH2)n-O(CH2)o-O(CH2)p-O(CH2)q-O(CH2)r-O-、
-N(R)-(CH2)m-O(CH2)n-O(CH2)o-O(CH2)p-O(CH2)q-O(CH2)r-O-、
-(CH2)m-O(CH2)n-O(CH2)o-O(CH2)p-O(CH2)q-O(CH2)r-O-、
-(CH2)m-O(CH2)n-O(CH2)o-O(CH2)p-O(CH2)q-O(CH2)r-OCH2-、


リンカーのm、n、o、p、q、およびrは、独立して、0、1、2、3、4、5、6、7、8、9、10、11、12、13、14、15、16、17、18、19、20であり、
数がゼロの場合、N-O結合またはO-O結合は存在しない。
リンカーのRは、H、メチルおよびエチルであり、
リンカーのXは、HおよびFであり、


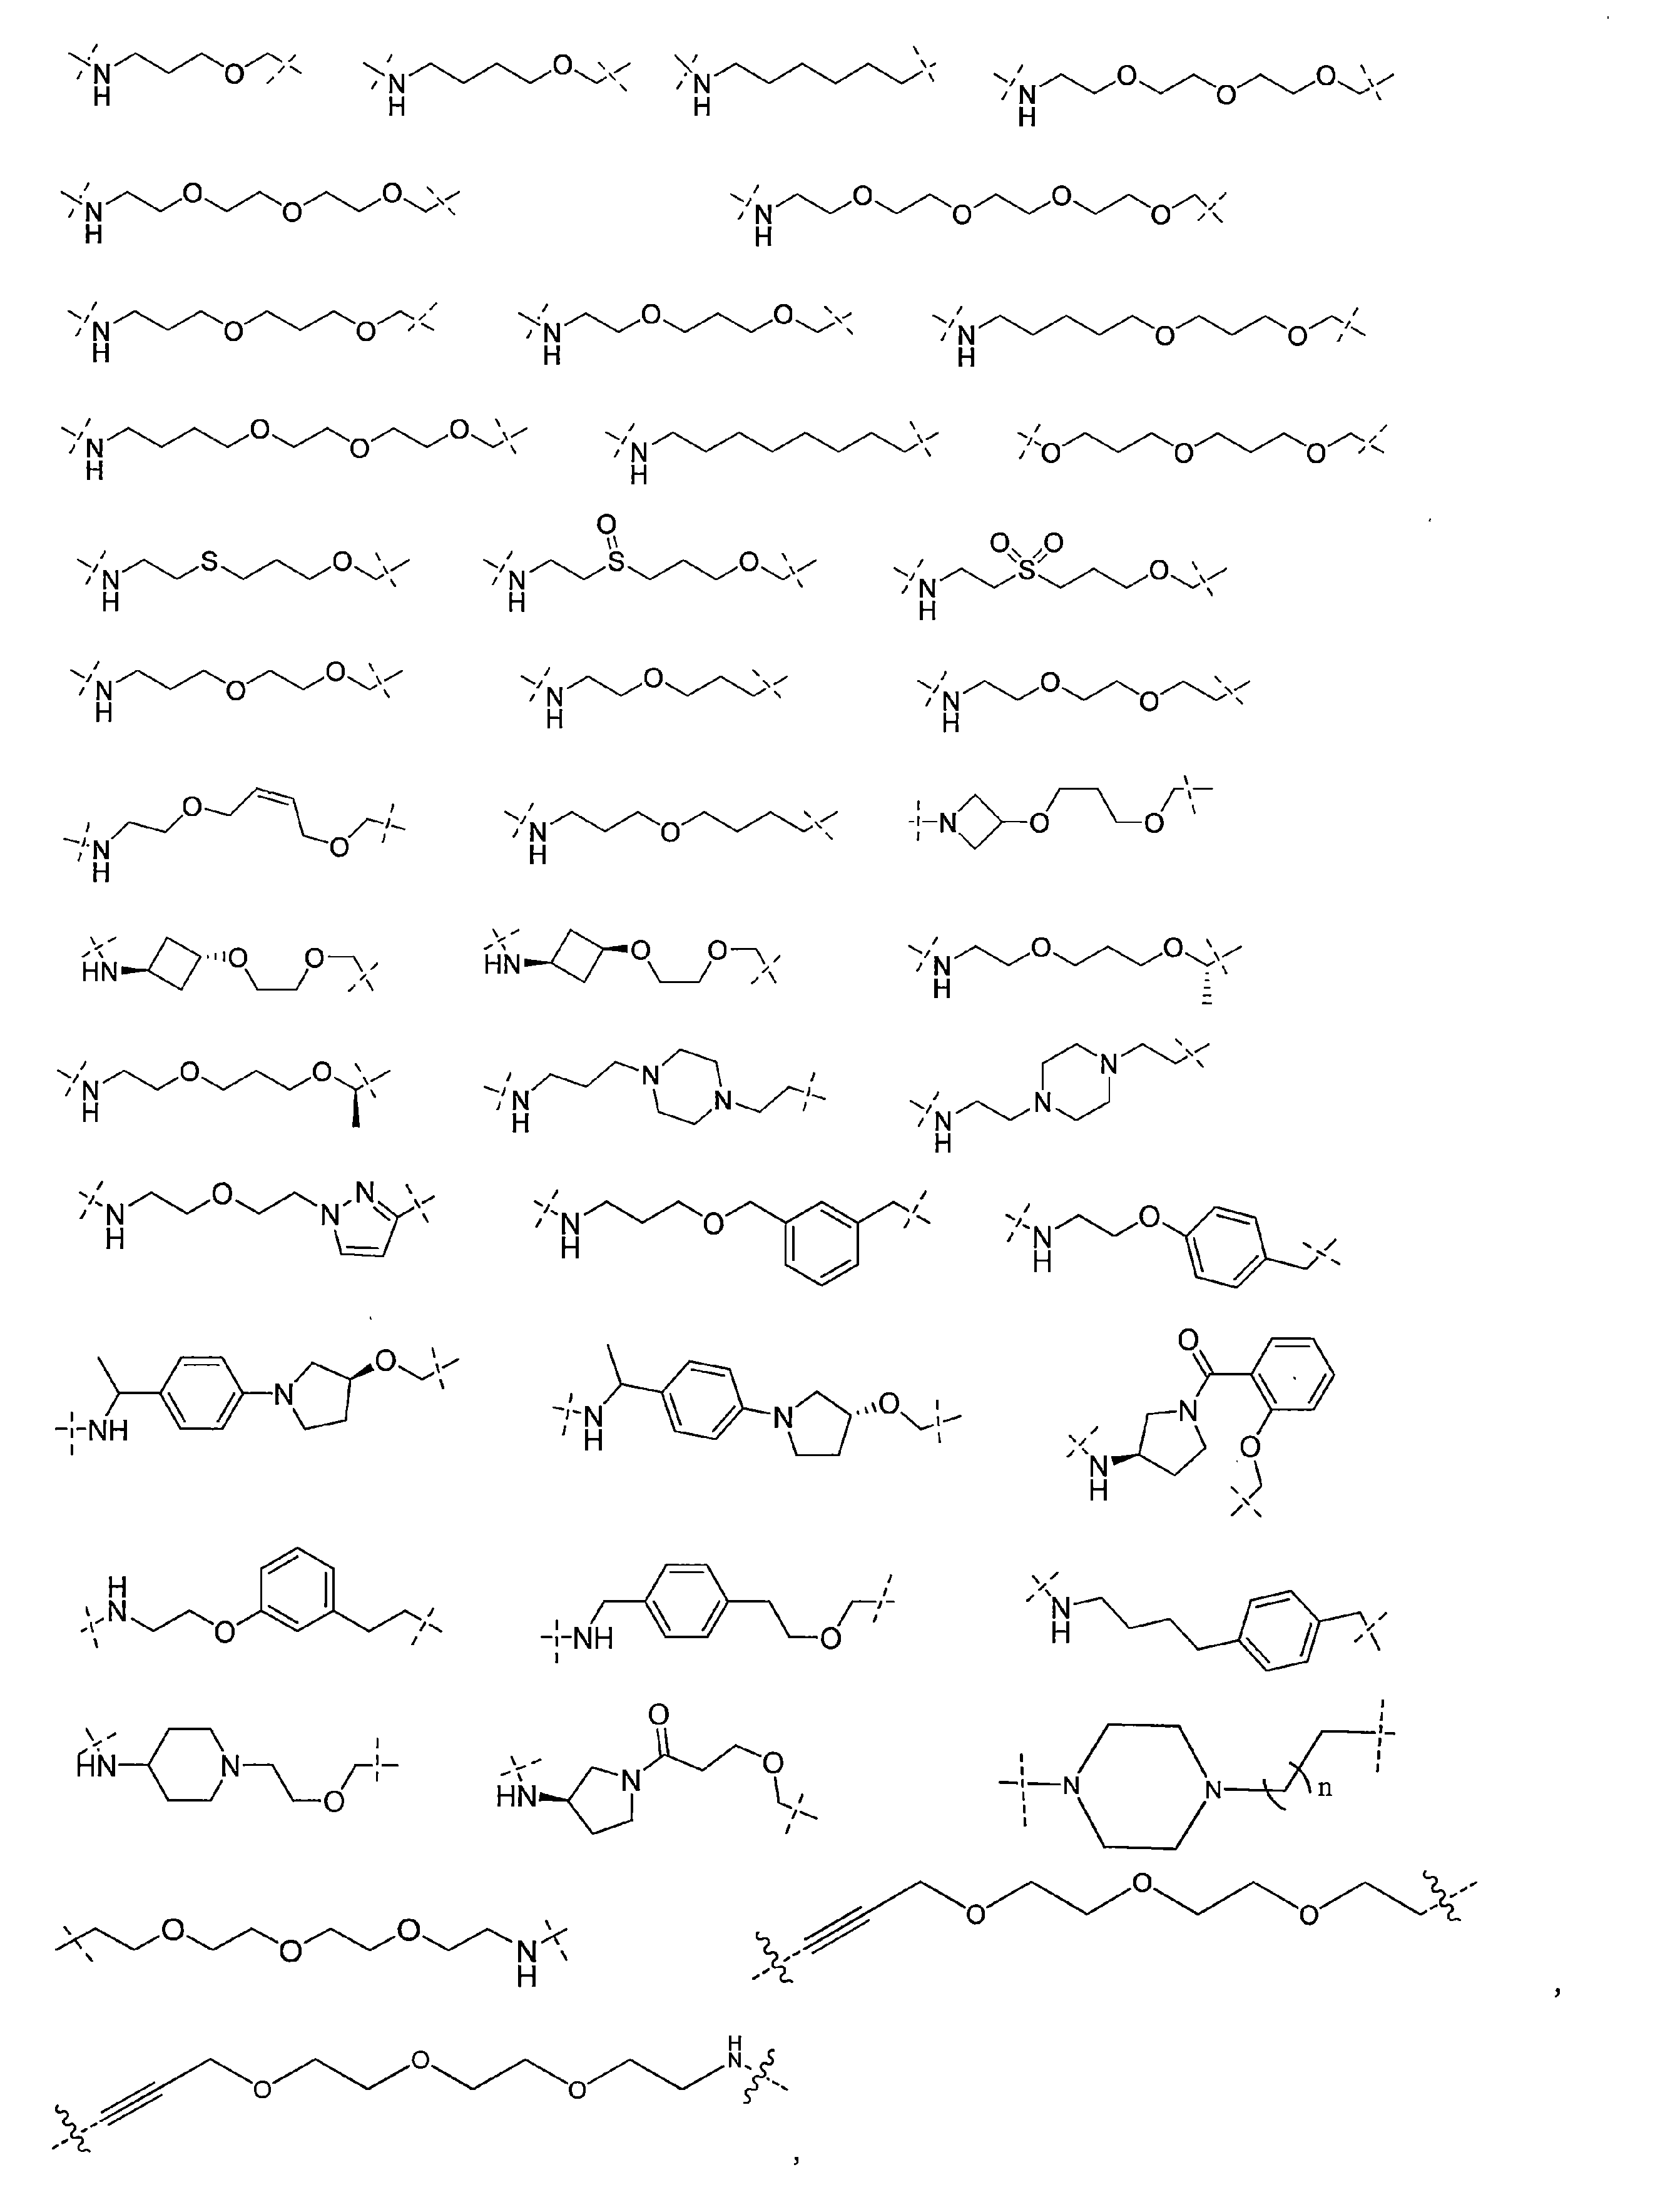


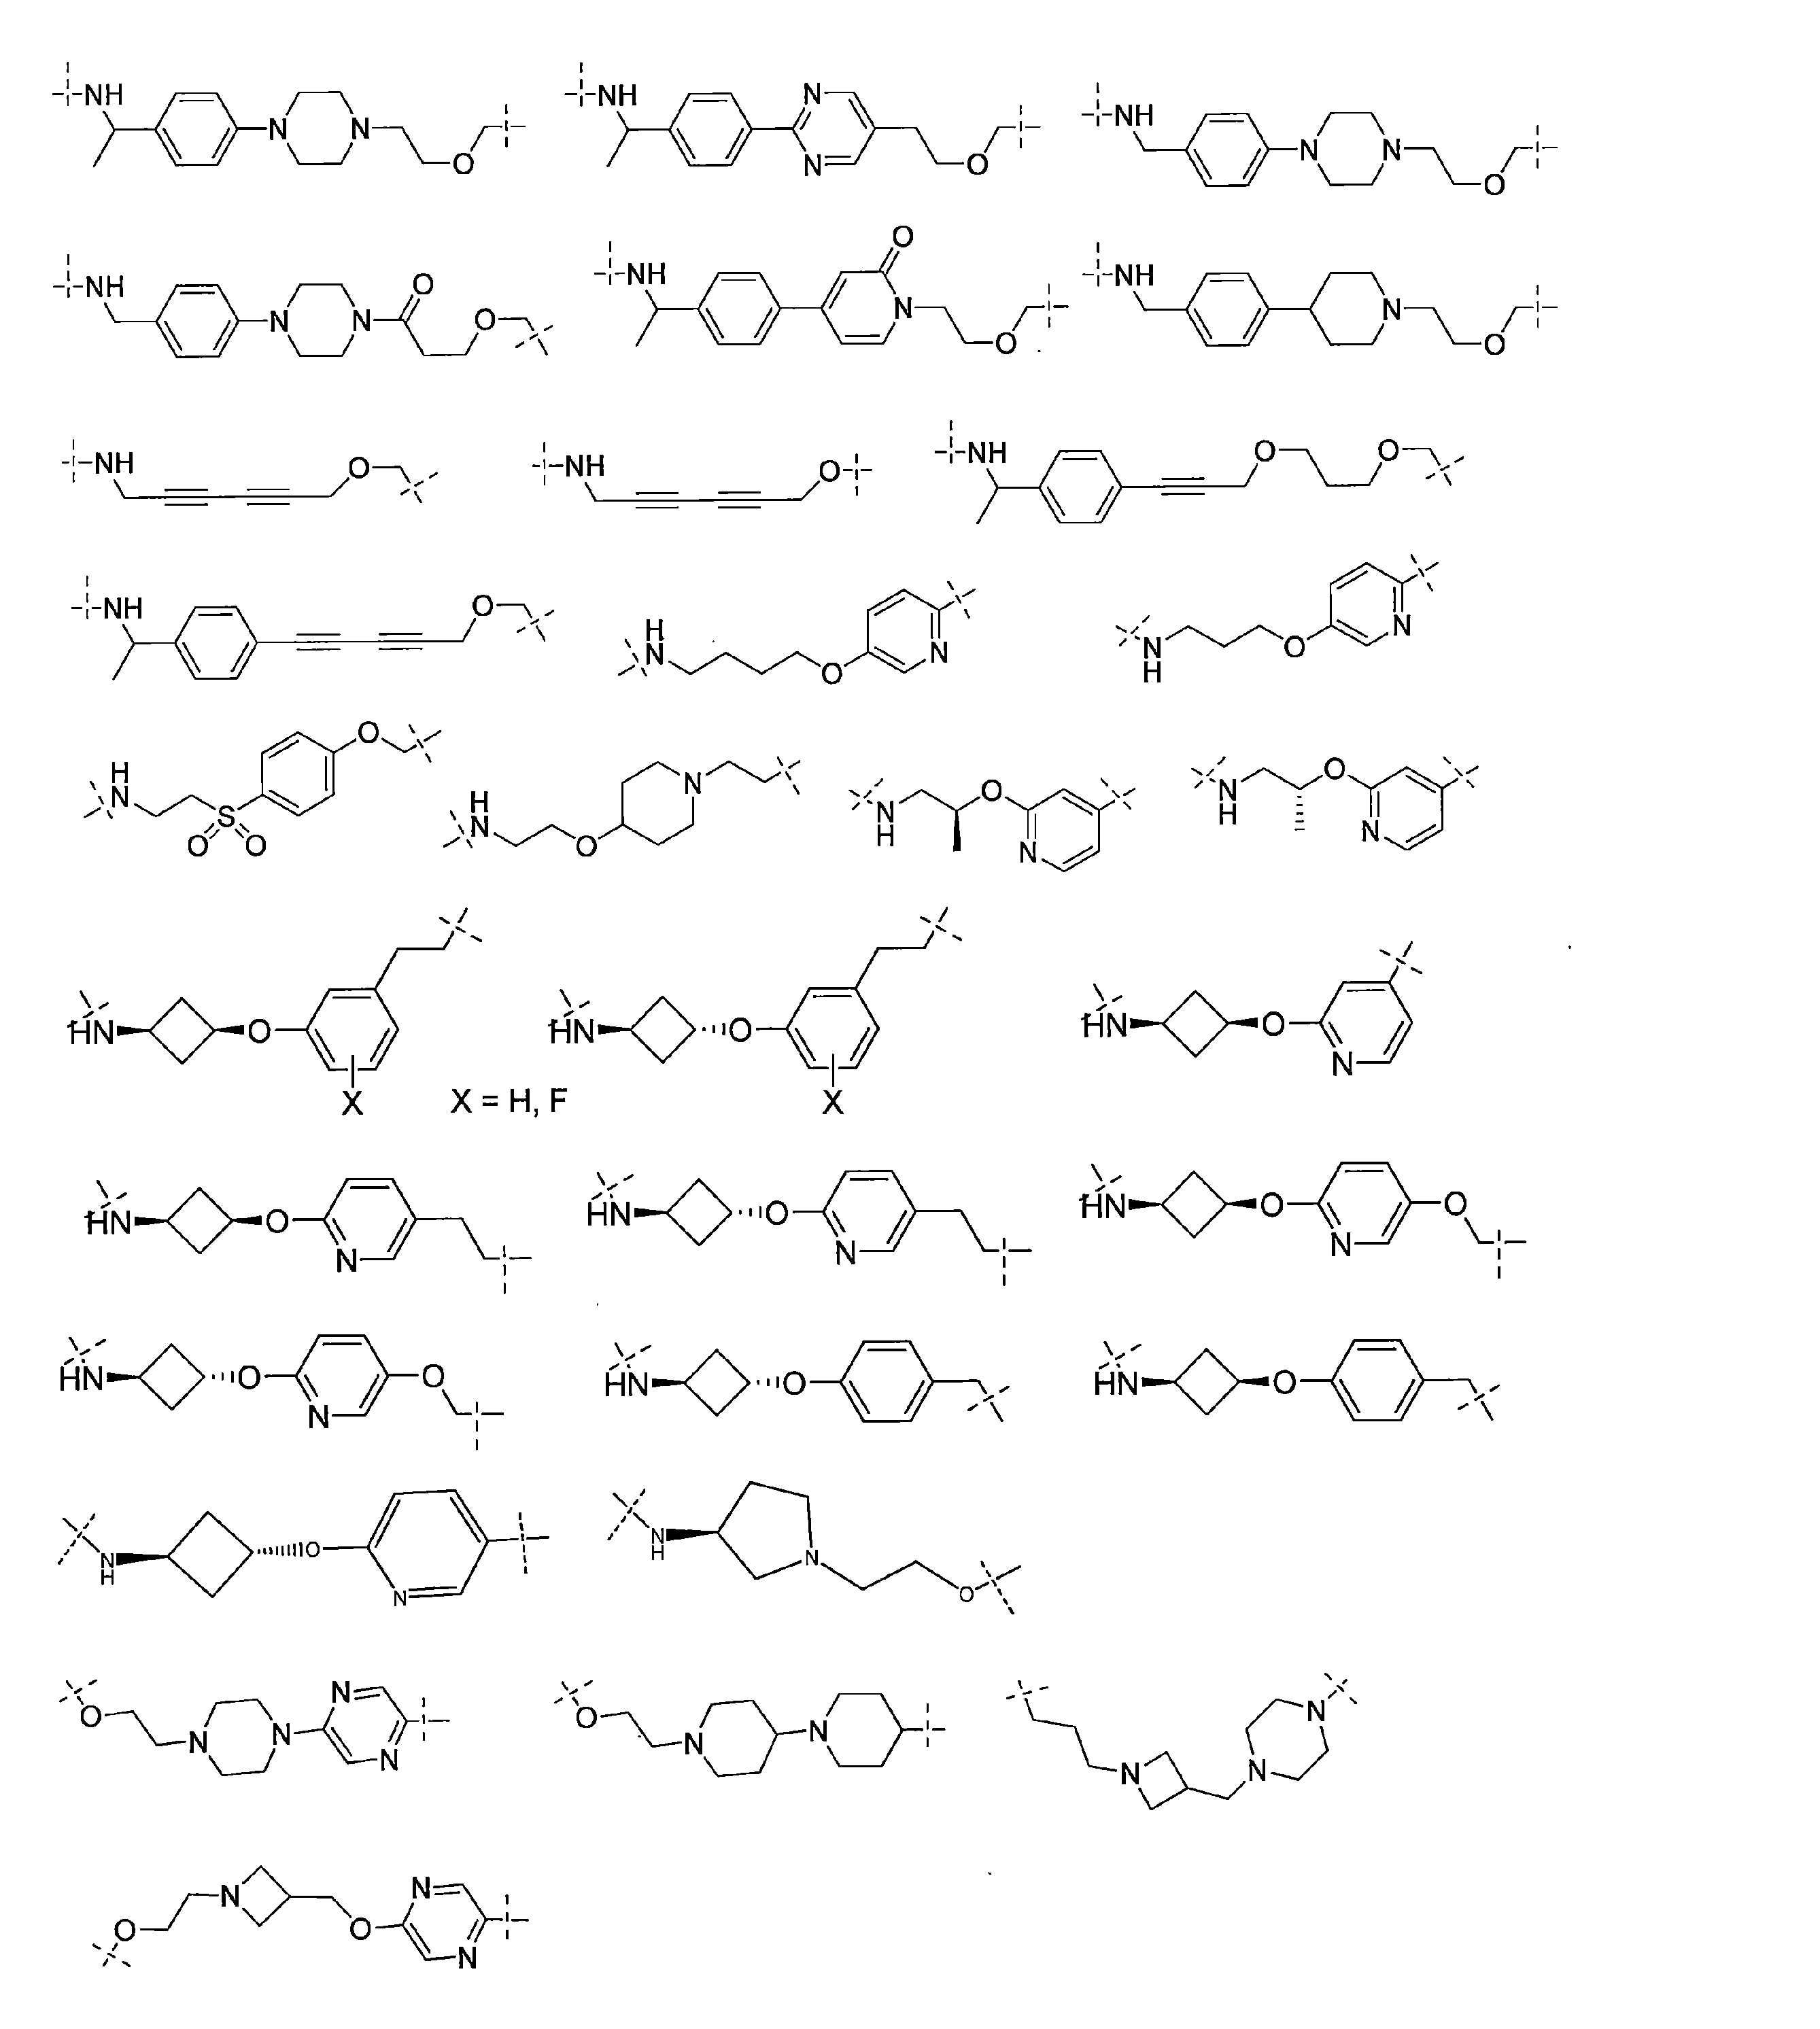

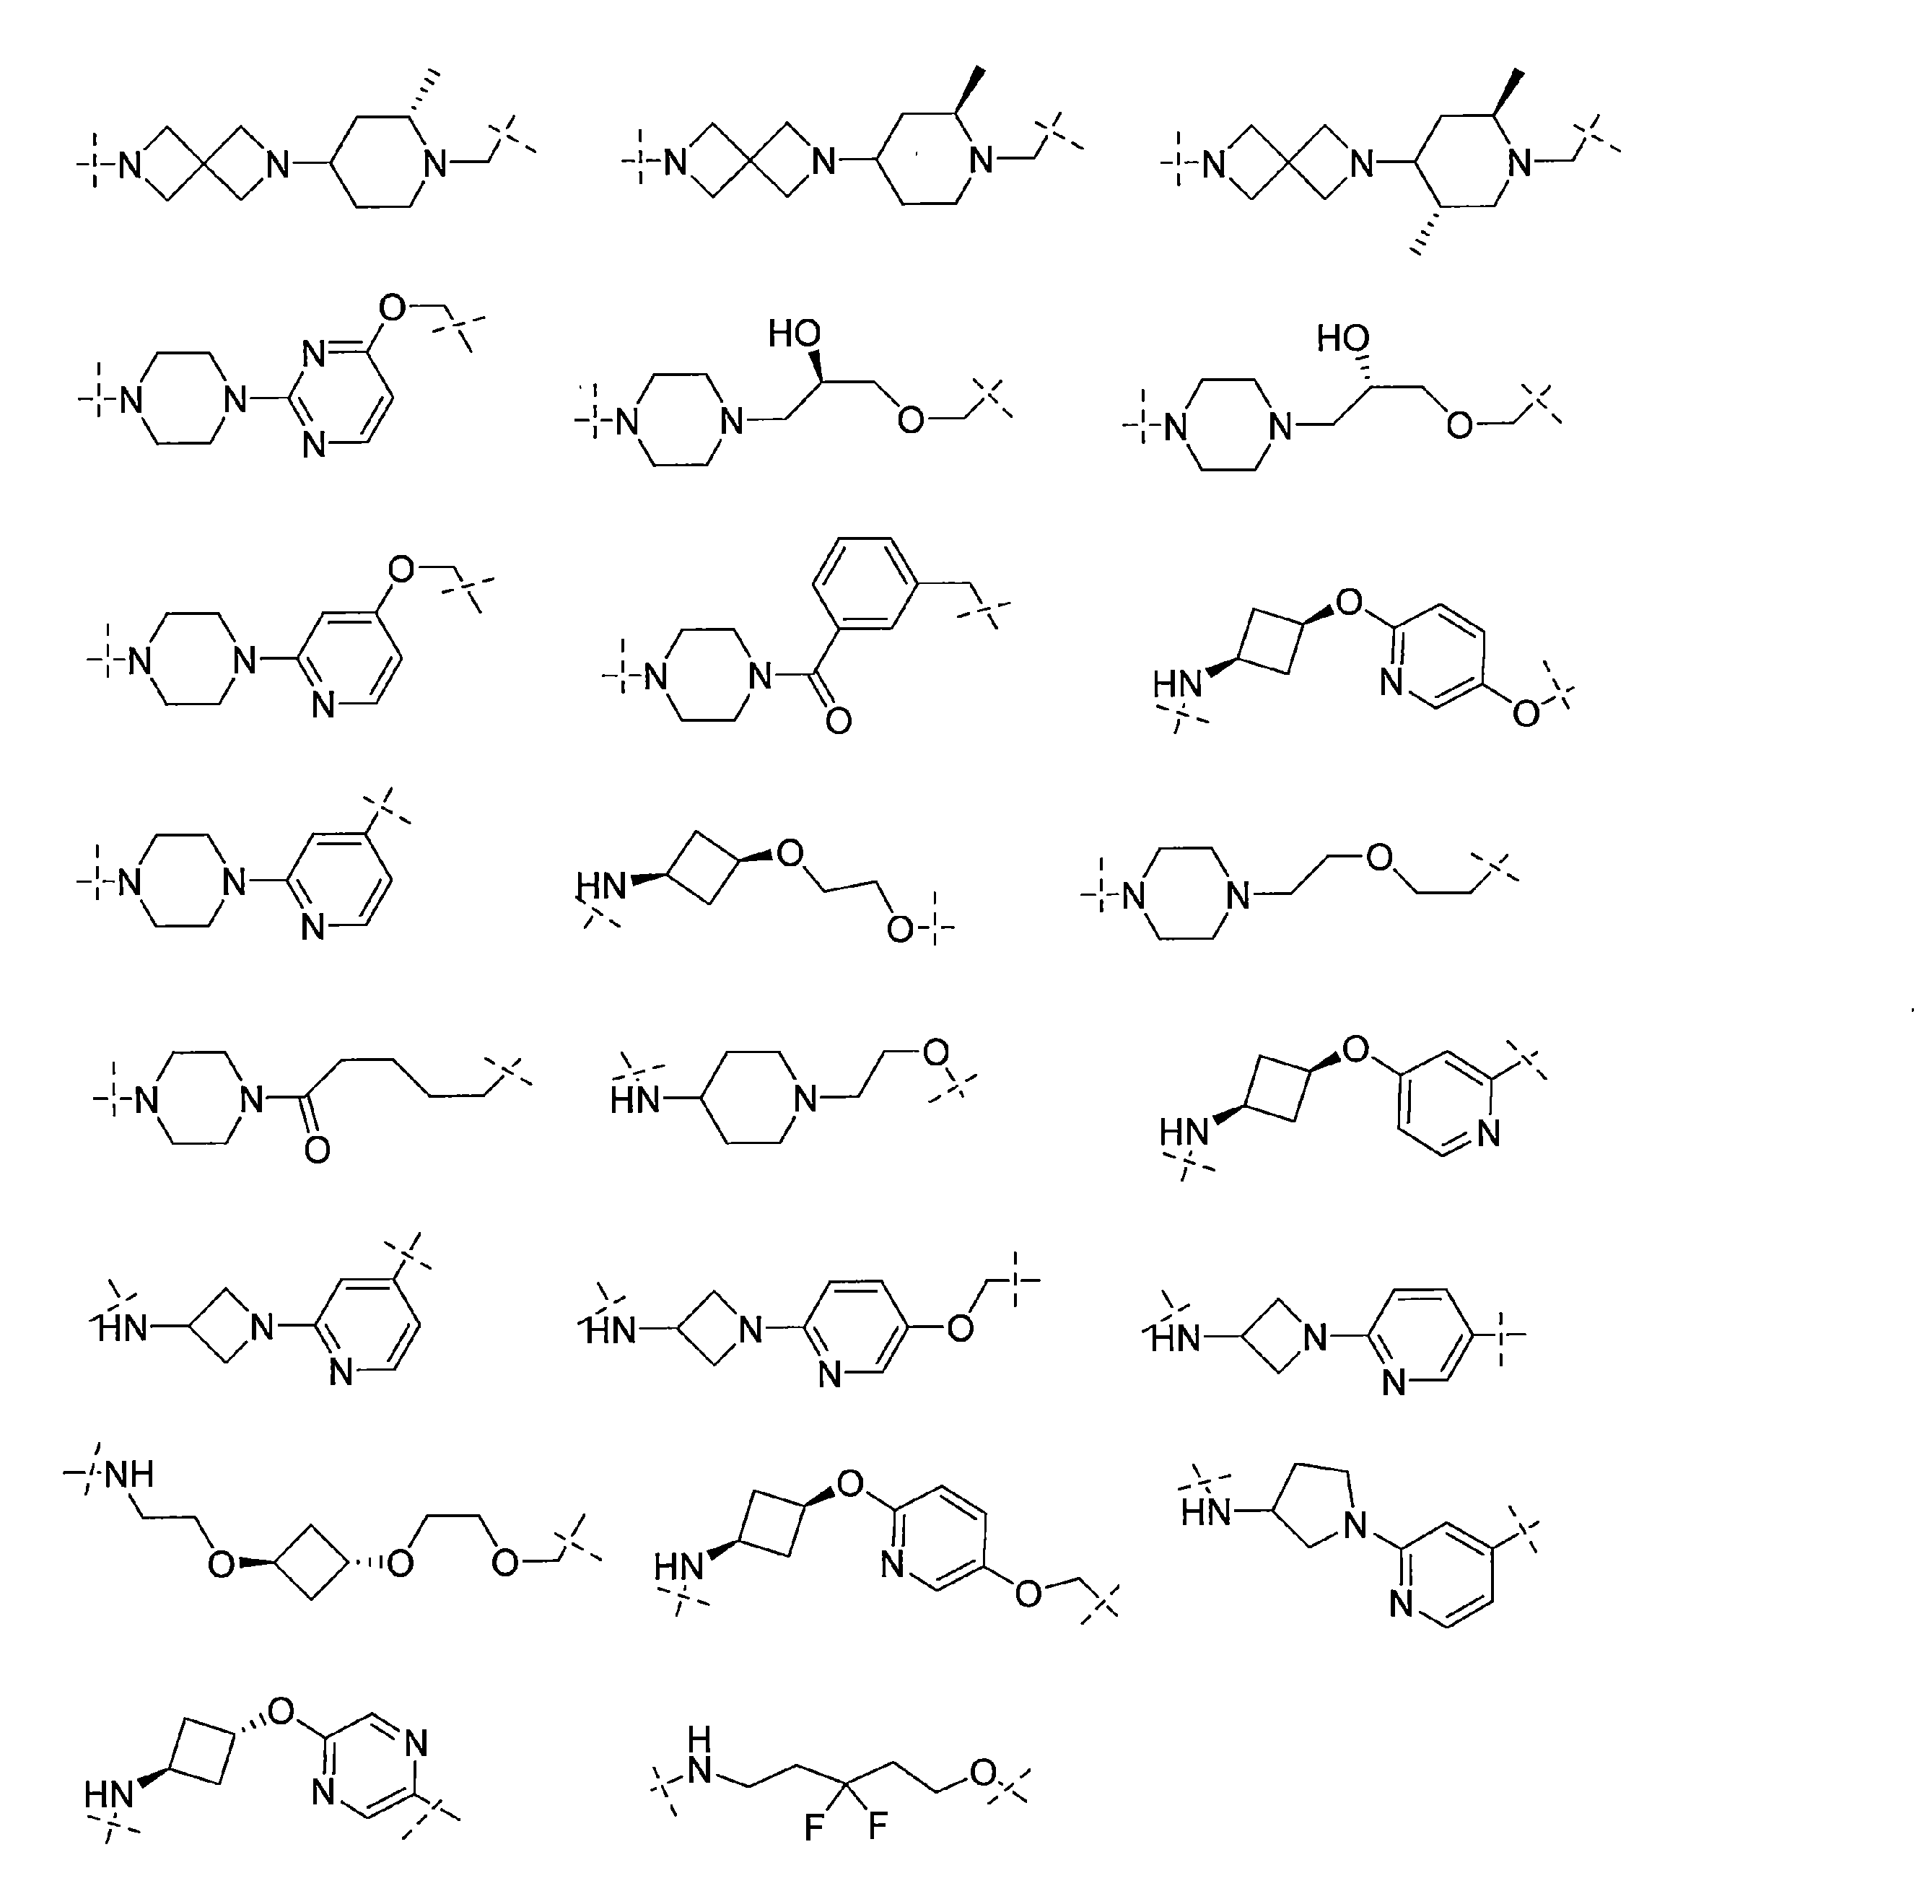
—N(R)—(CH2)m-O(CH2)n-O(CH2)o-O(CH2)p-O(CH2)q-O(CH2)r-OCH2-,
—O—(CH2)m—O(CH2)n—O(CH2)o—O(CH2)p—O(CH2)q—O(CH2)r—OCH2—,
—O—(CH2)m—O(CH2)n—O(CH2)o—O(CH2)p—O(CH2)q—O(CH2)r—O—,
—N(R)—(CH2)m-O(CH2)n-O(CH2)o-O(CH2)p-O(CH2)q-O(CH2)r-O-,
-(CH2)m-O(CH2)n-O(CH2)o-O(CH2)p-O(CH2)q-O(CH2)r-O-,
-(CH2)m-O(CH2)n-O(CH2)o-O(CH2)p-O(CH2)q-O(CH2)r-OCH2-,
m, n, o, p, q, and r of the linker are independently 0, 1, 2, 3, 4, 5, 6, 7, 8, 9, 10, 11, 12, 13, 14, 15, 16, 17, 18, 19, 20;
When the number is zero, there are no NO or OO bonds.
The linker R is H, methyl and ethyl;
X of the linker is H and F;
本明細書に記載される任意の態様または実施形態では、リンカー(L)の単位ALは、以下からなる群から選択され、
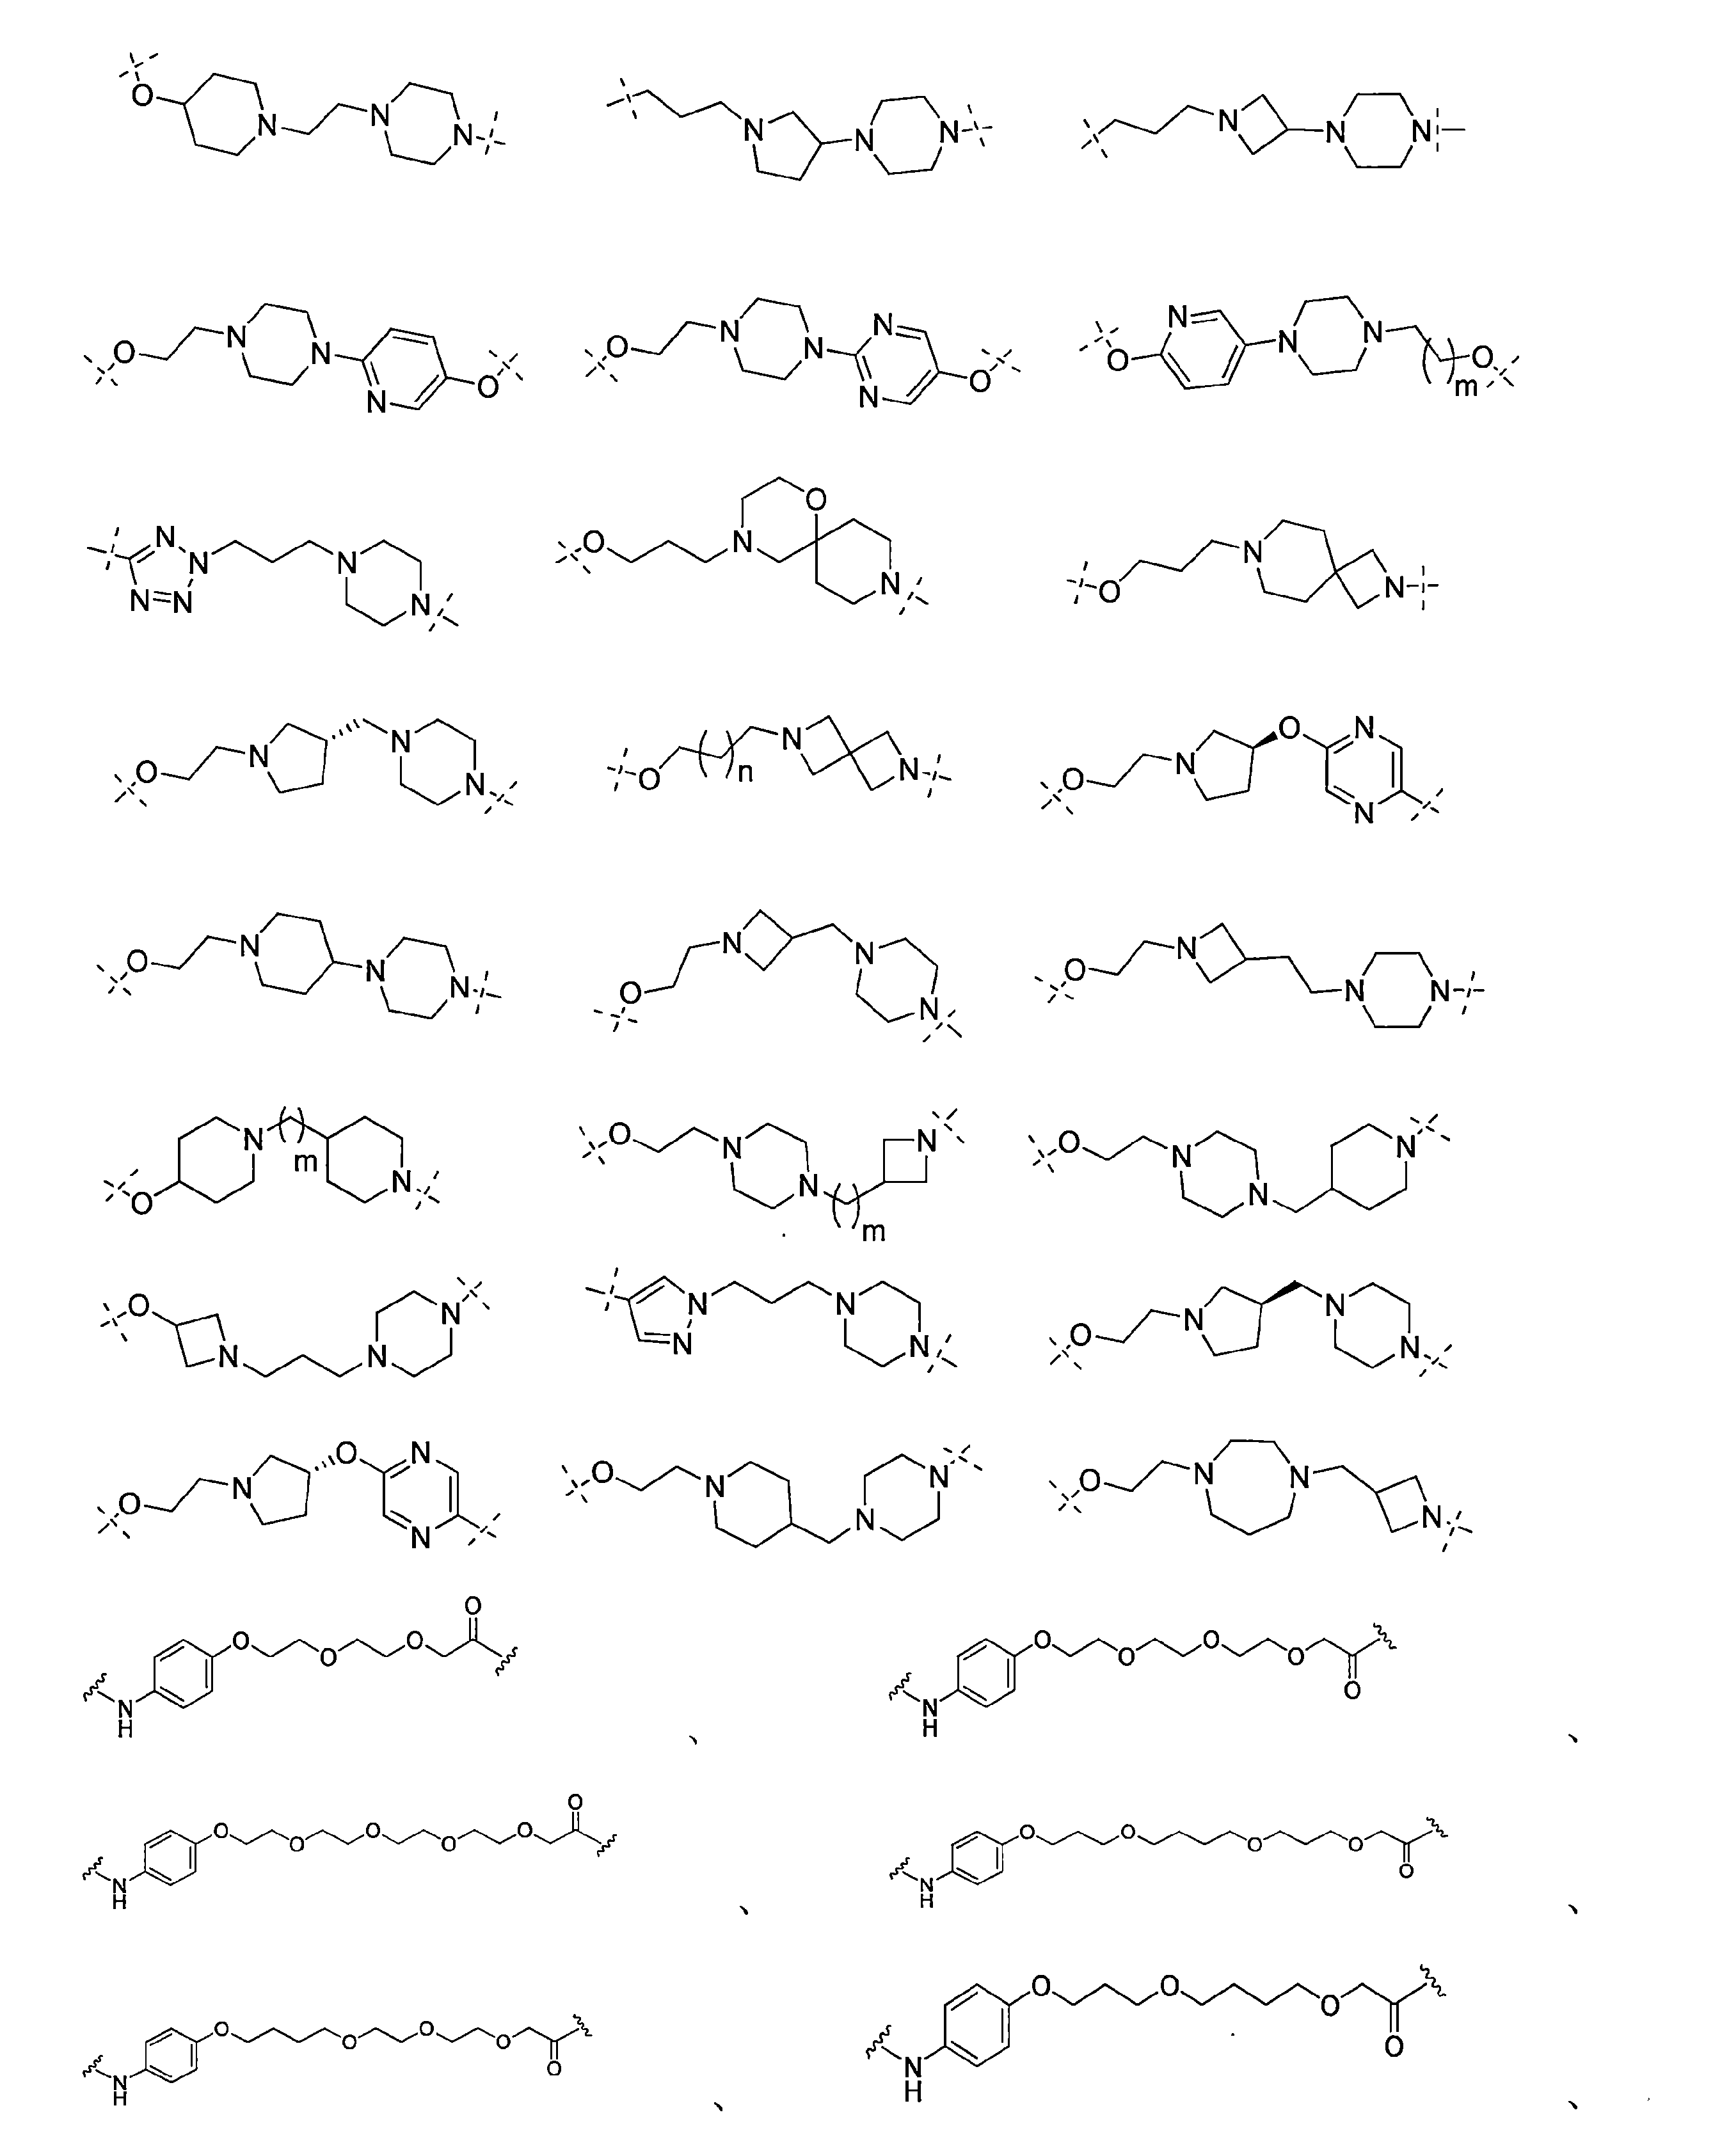


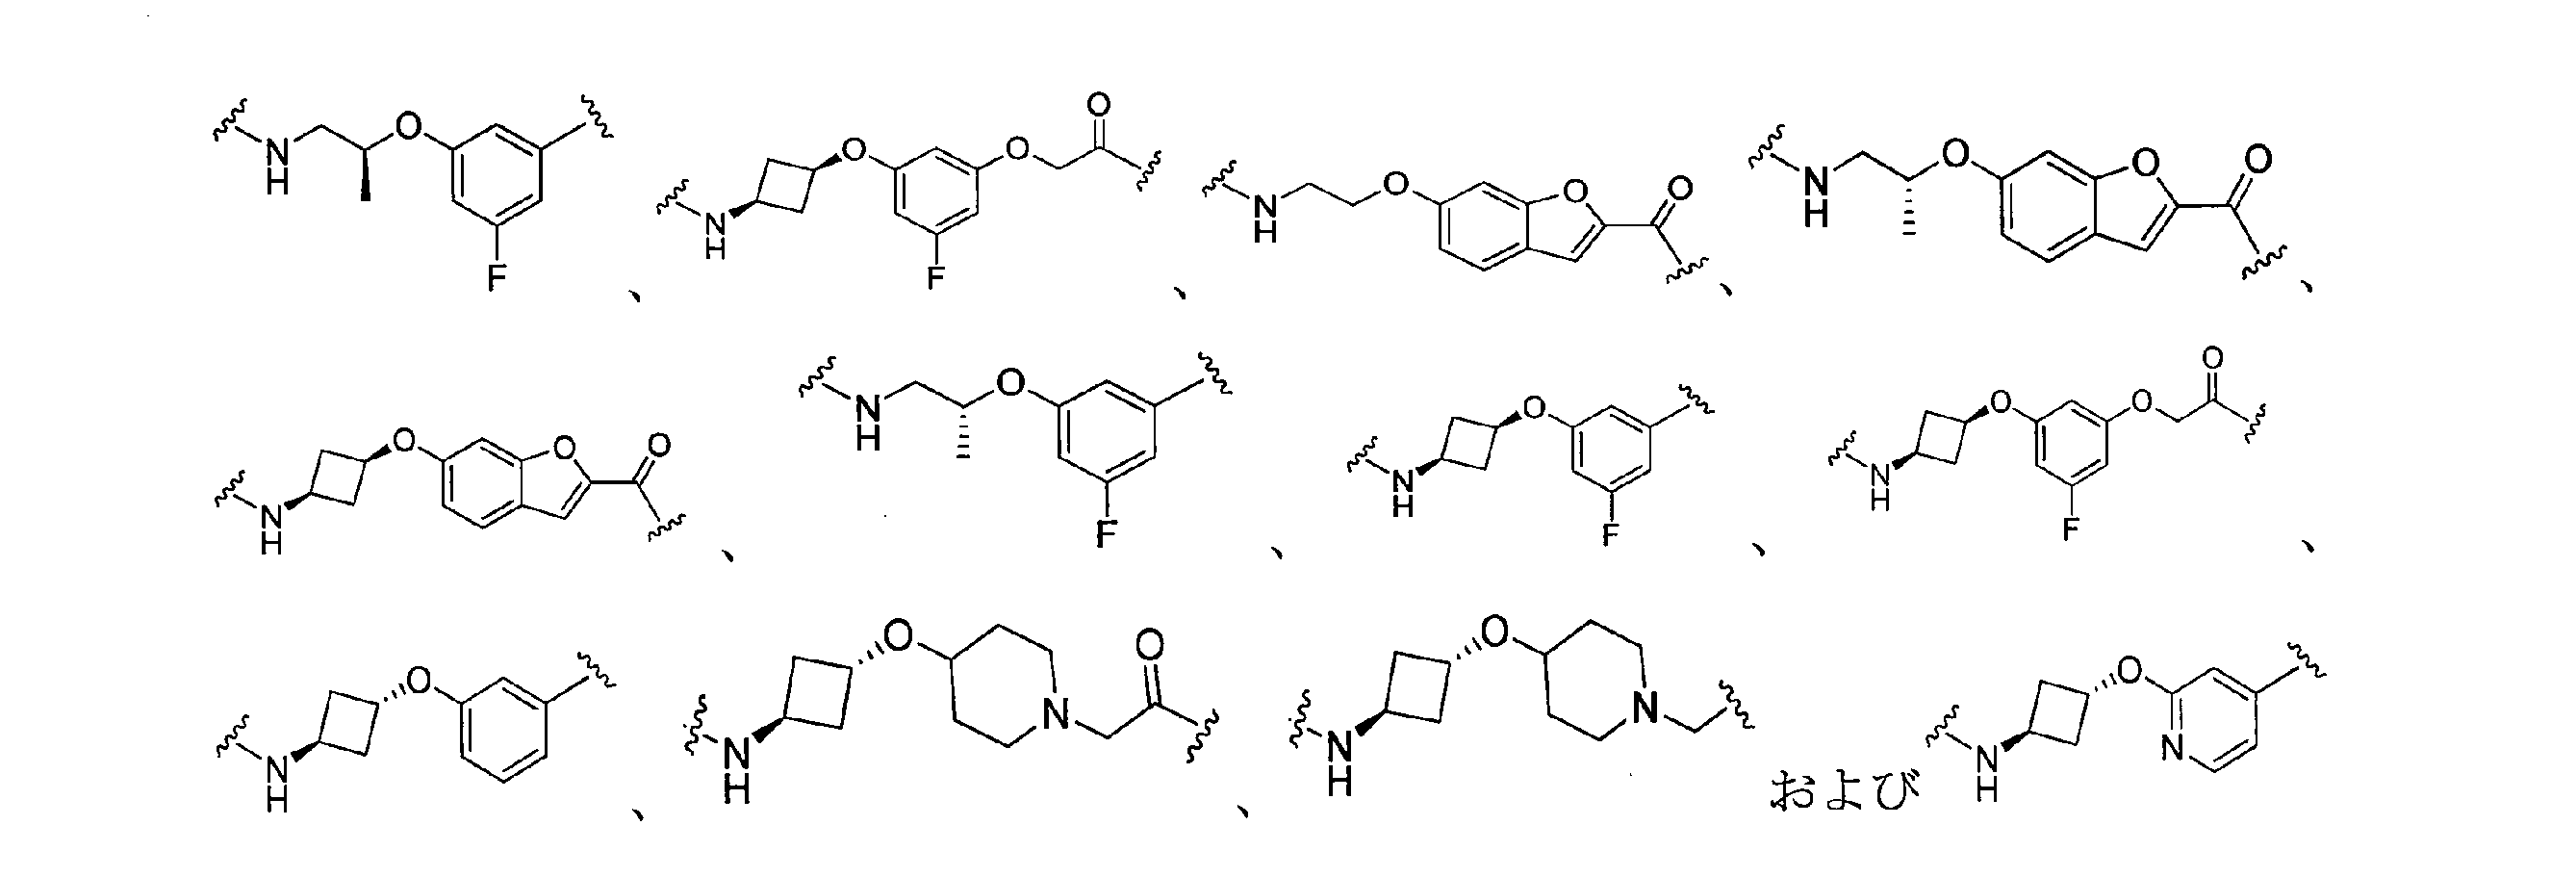
本明細書に記載される任意の態様または実施形態では、リンカー(L)の単位ALは、以下からなる群から選択され、


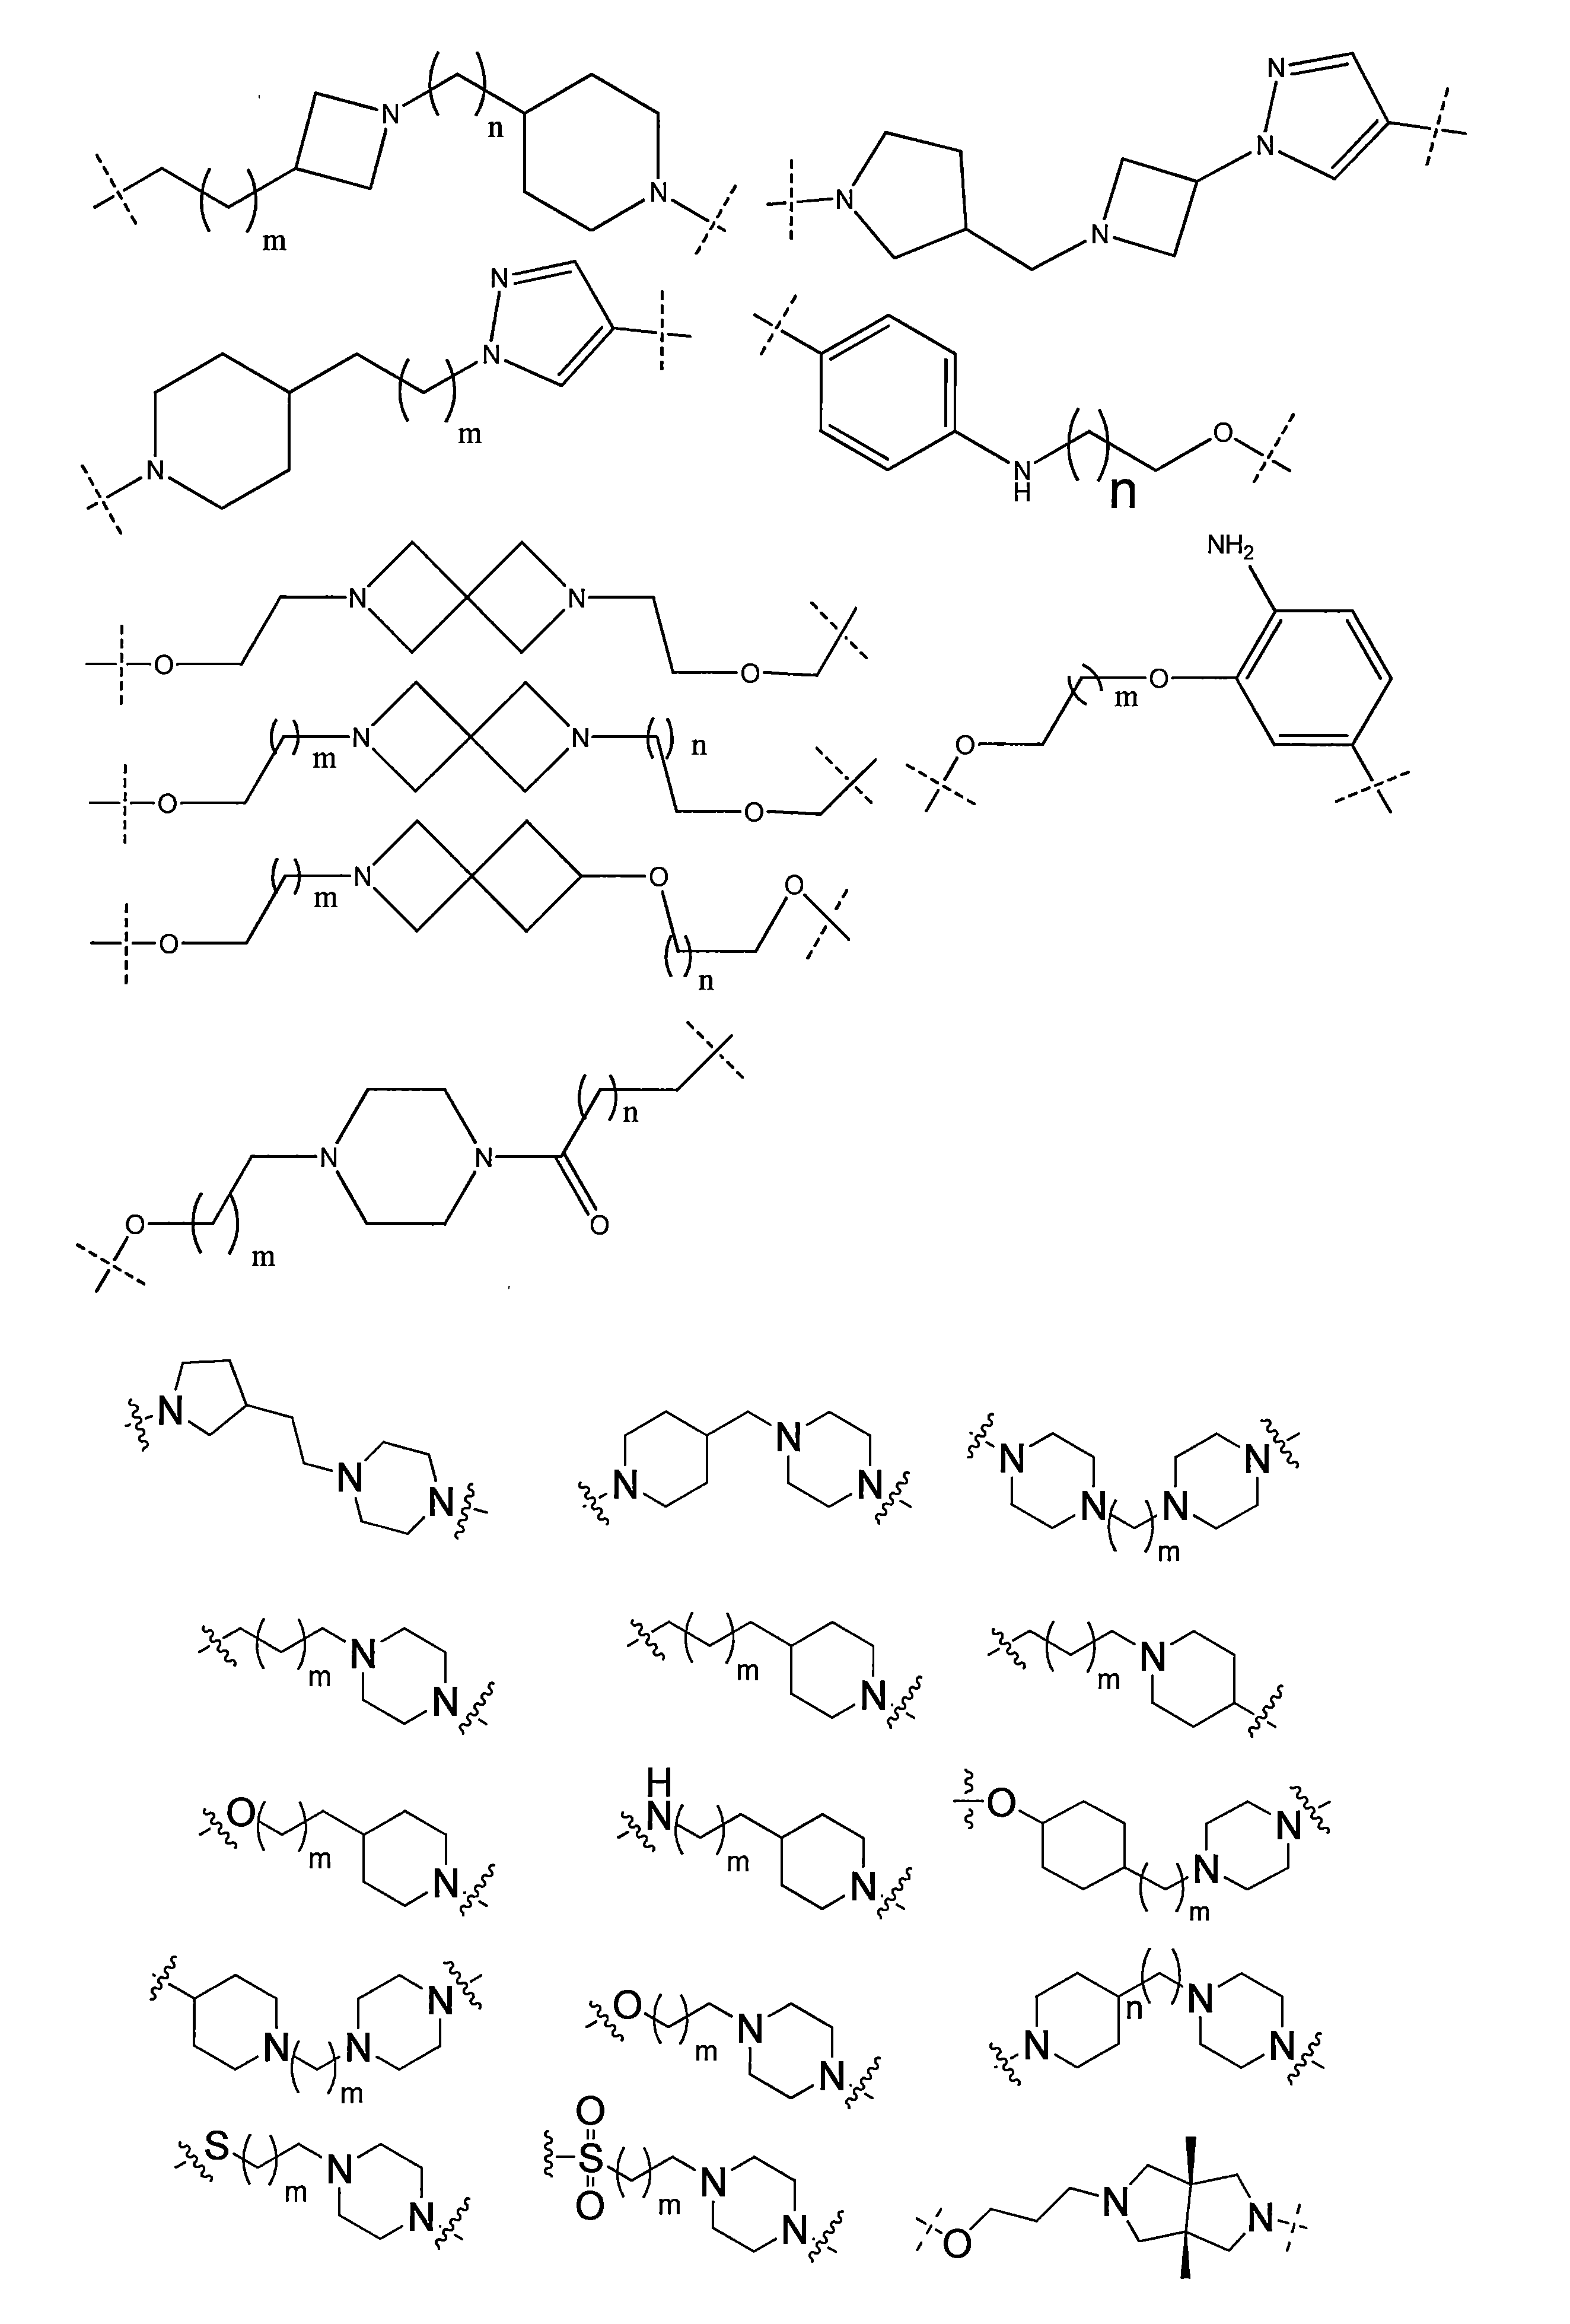

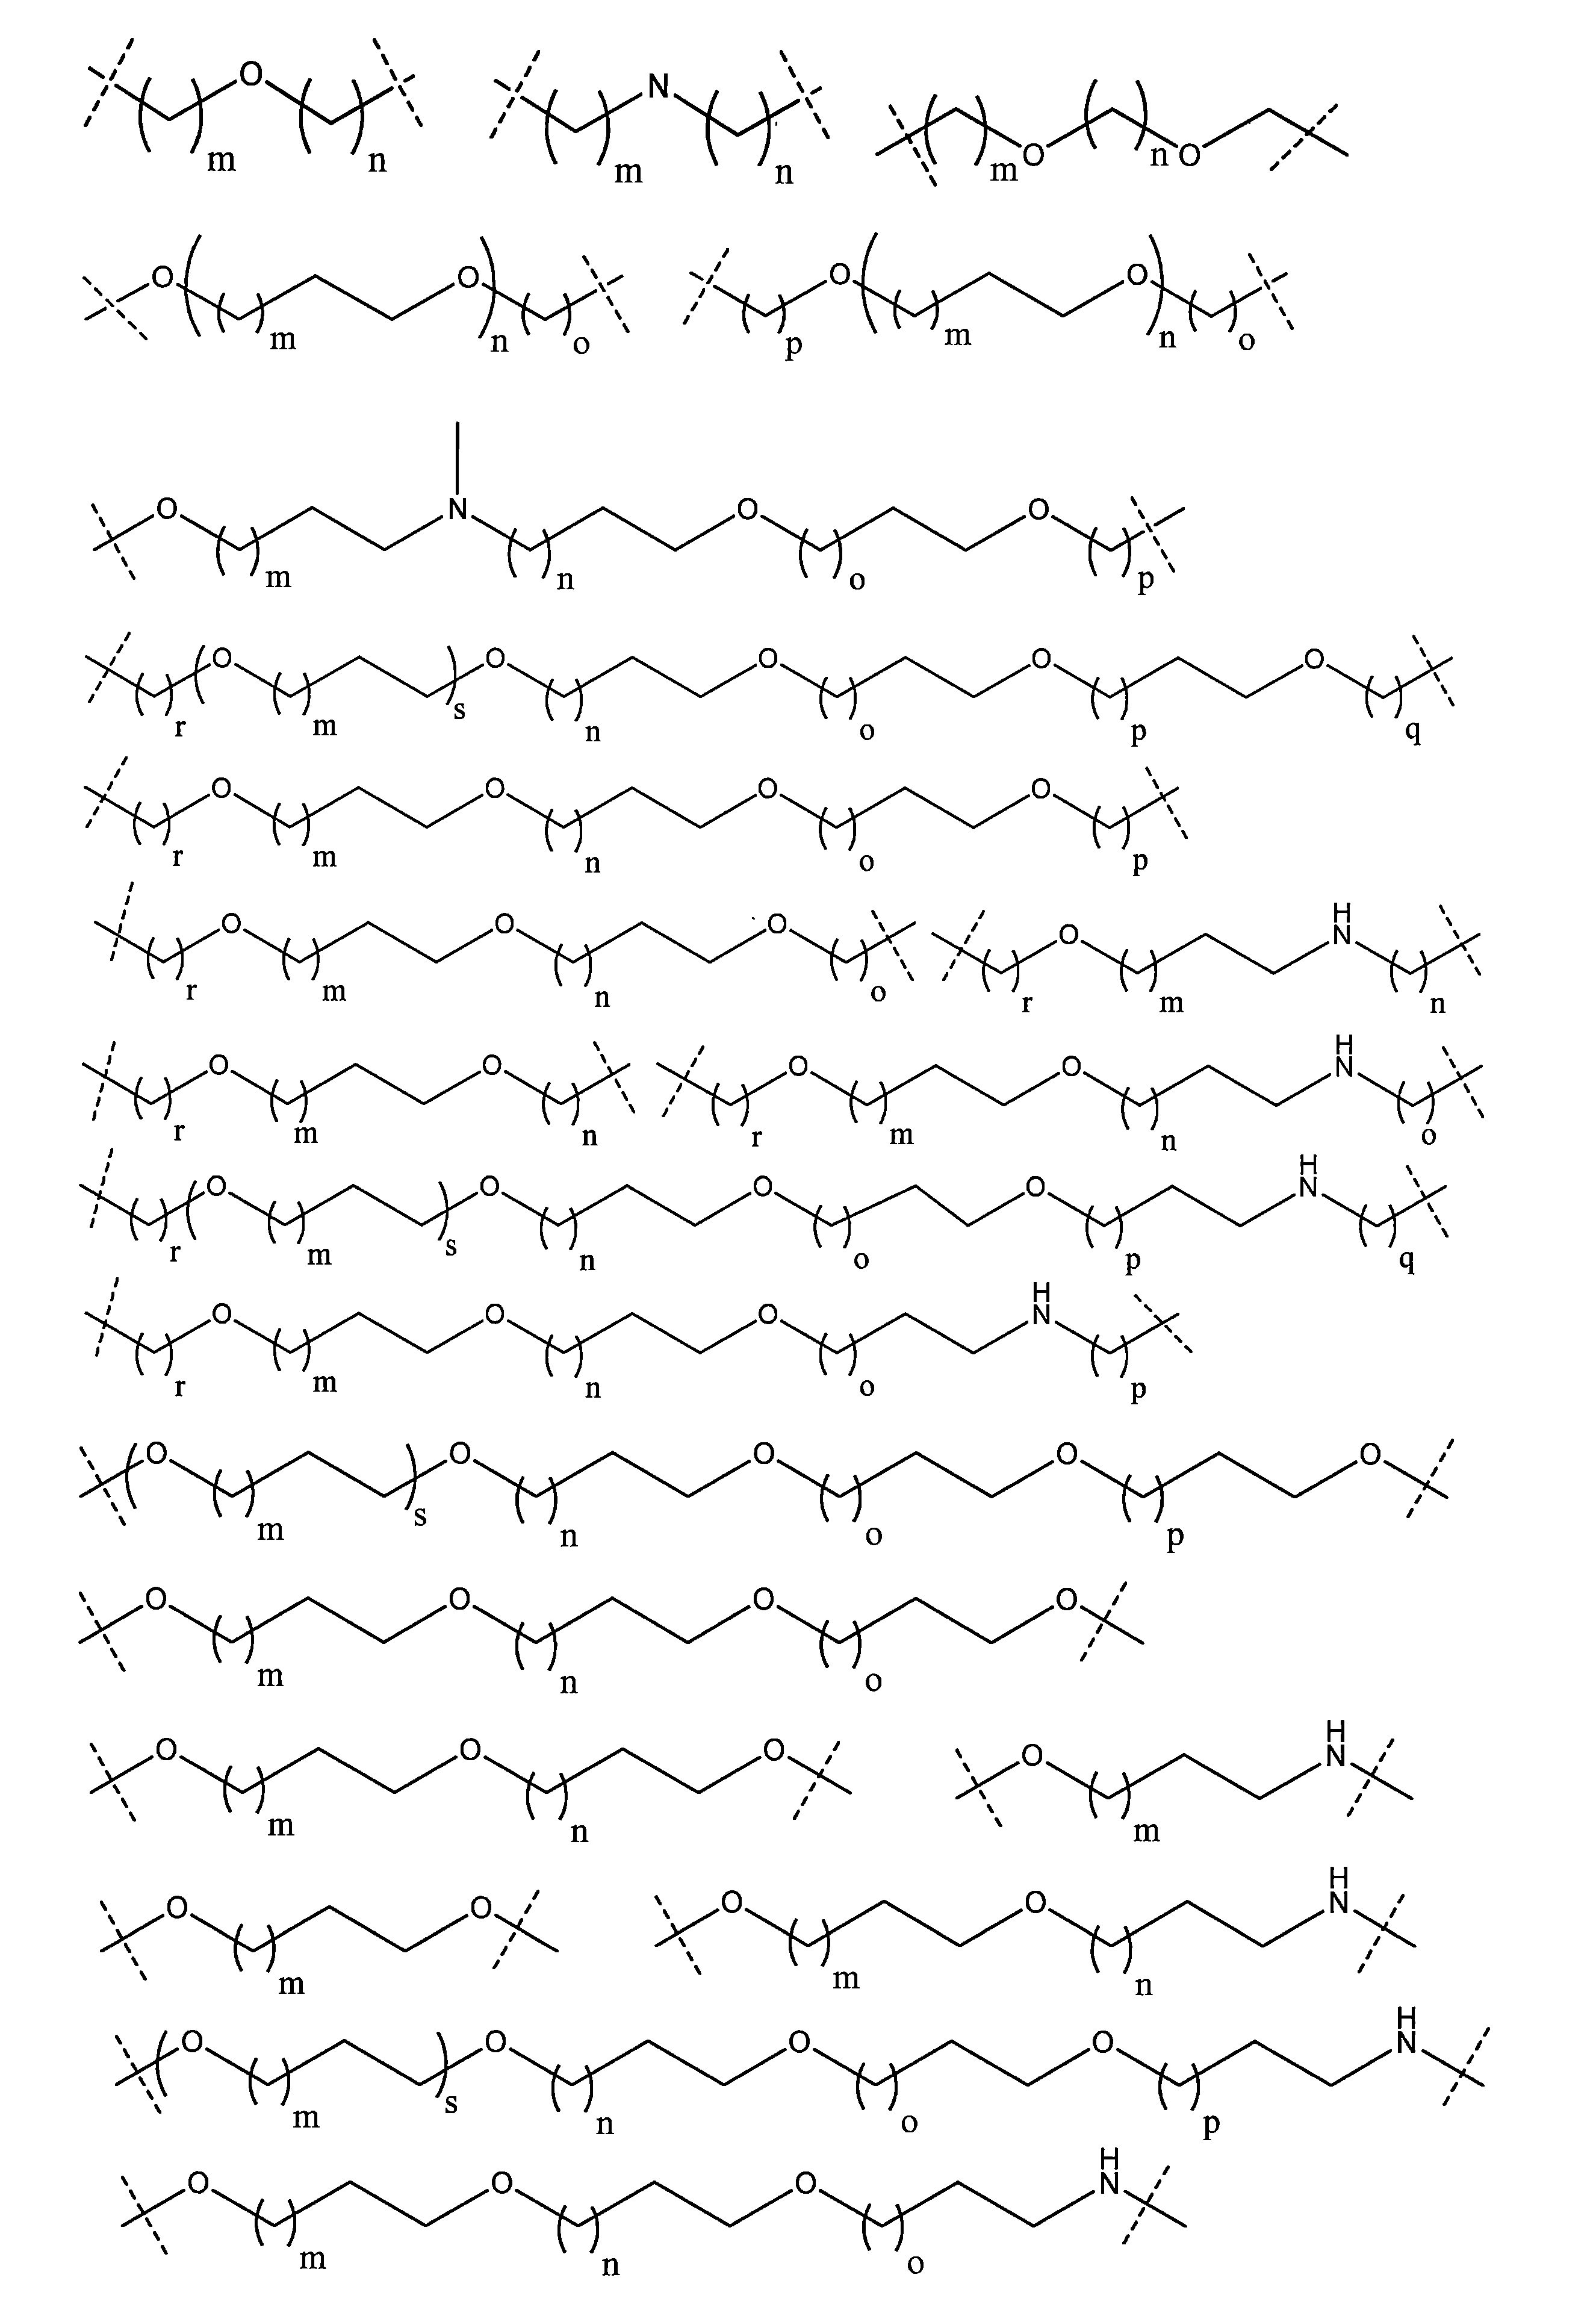


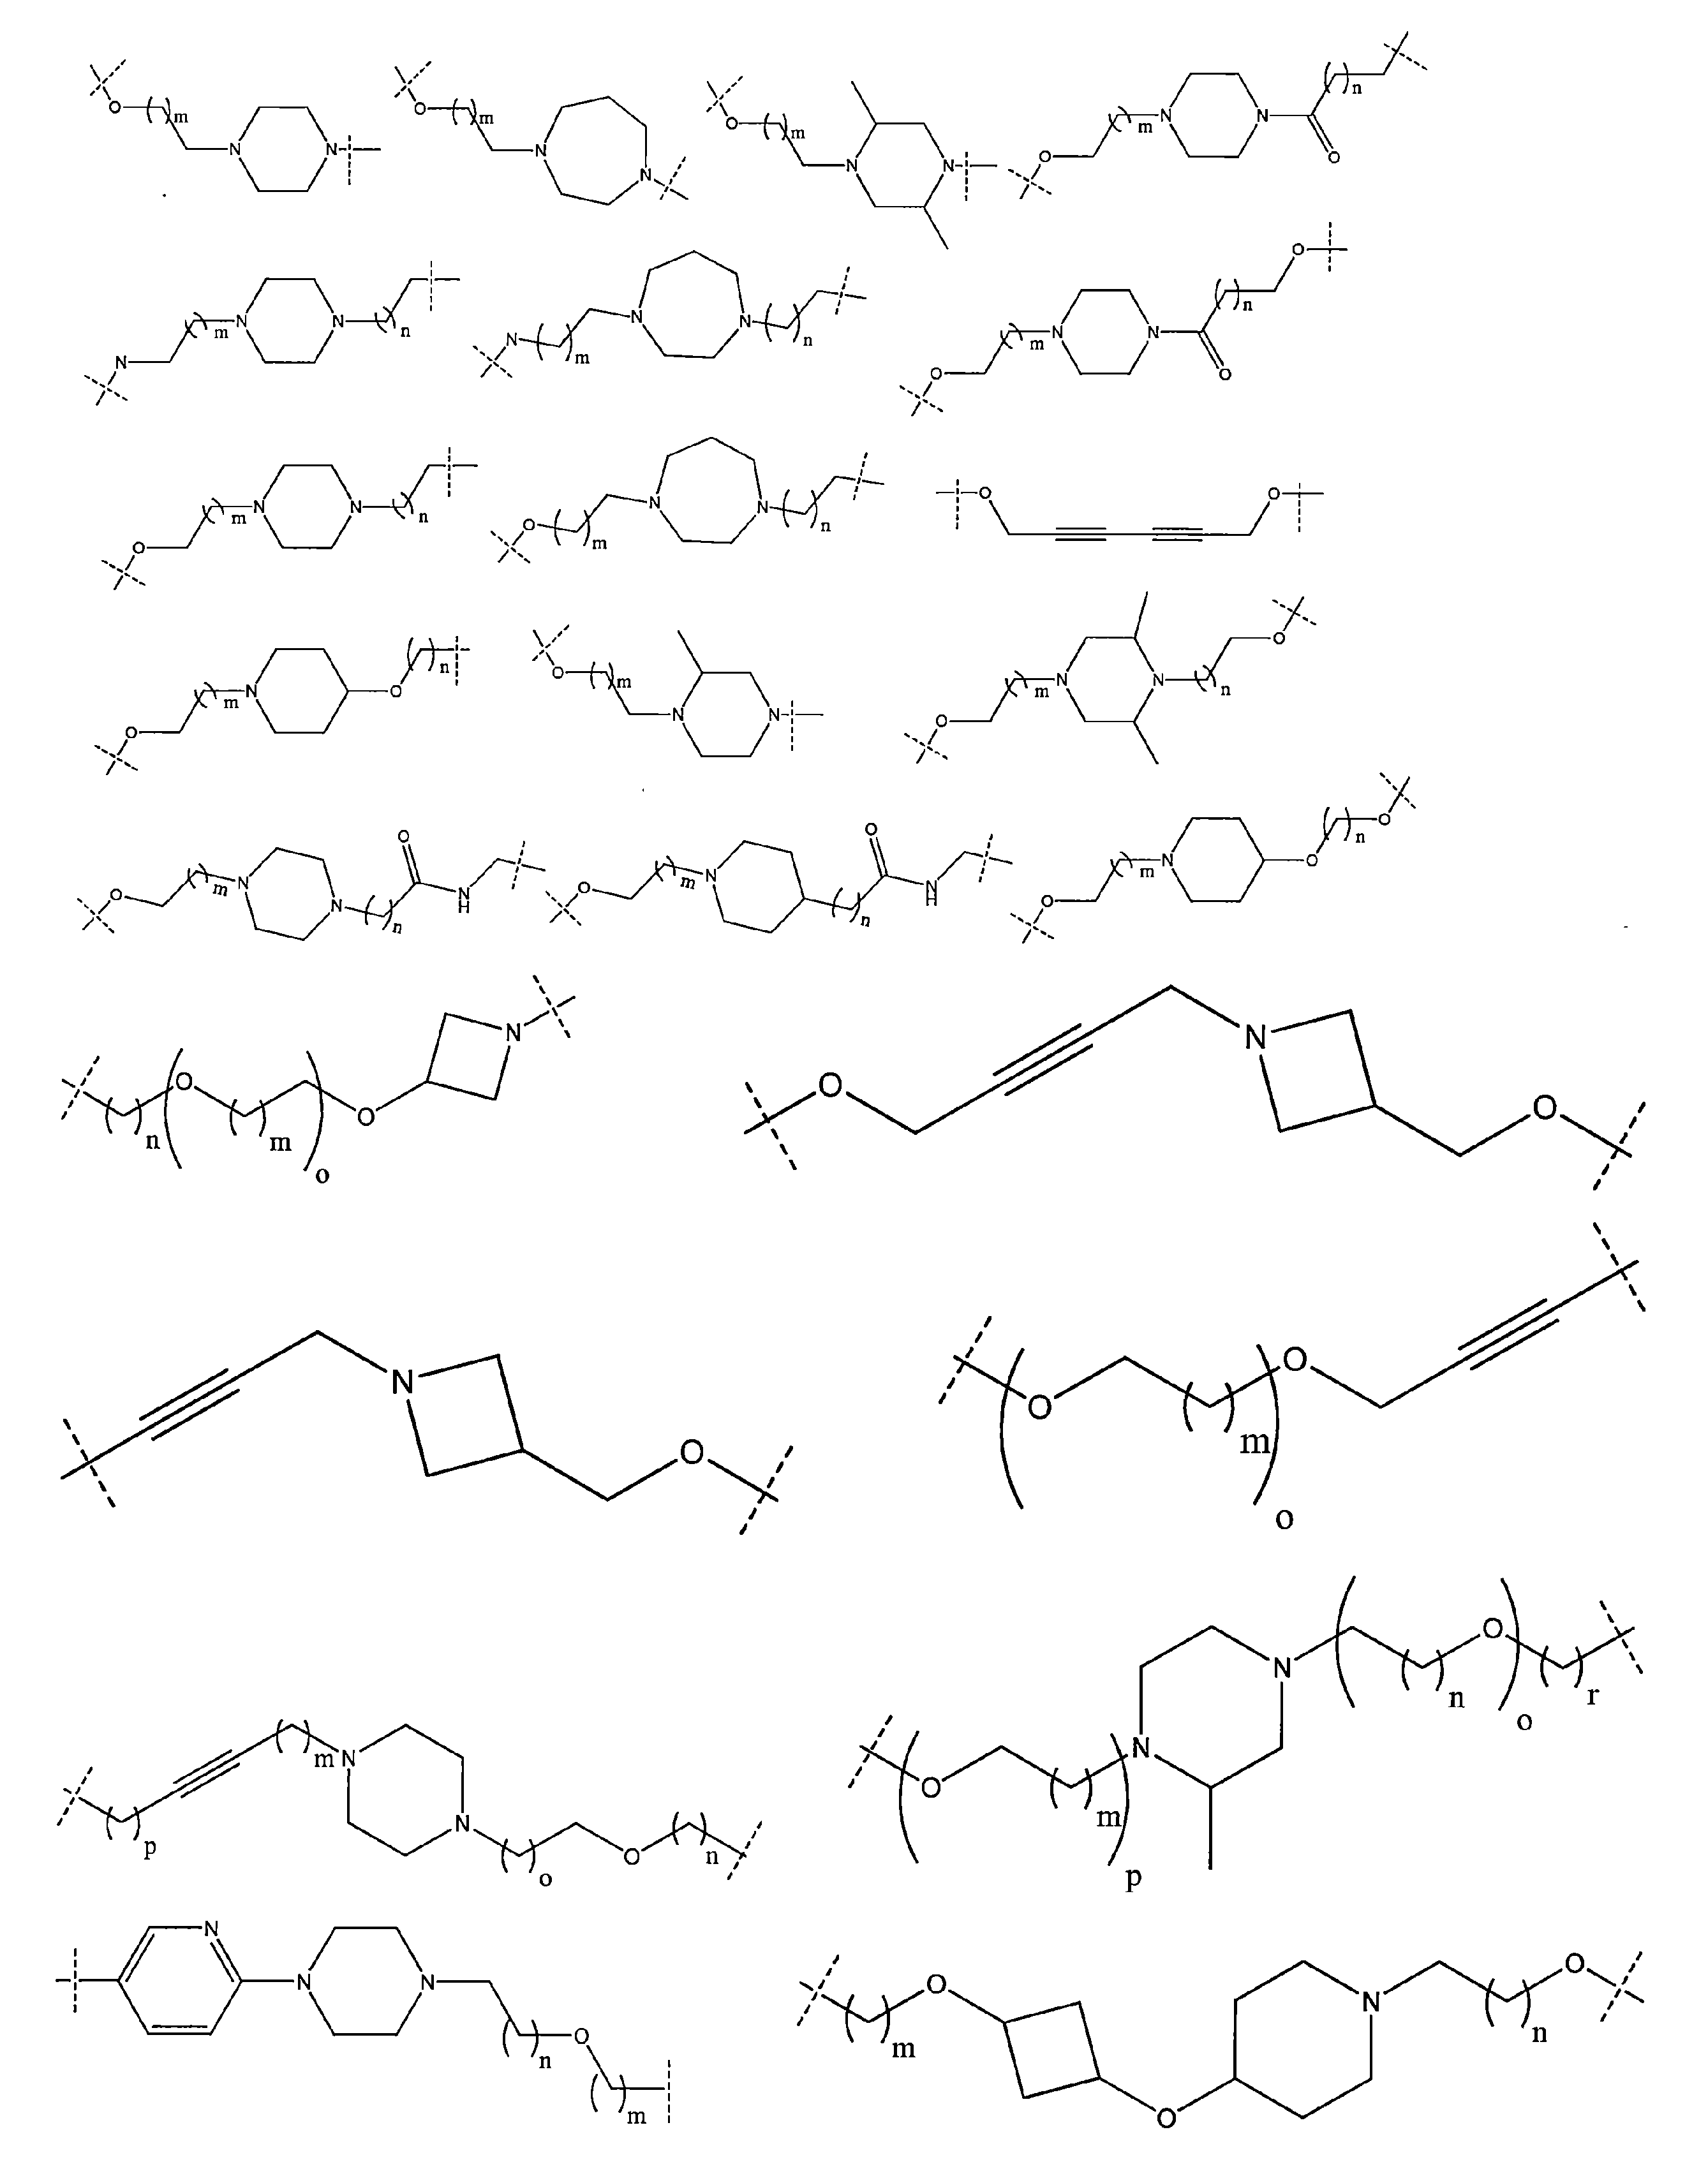


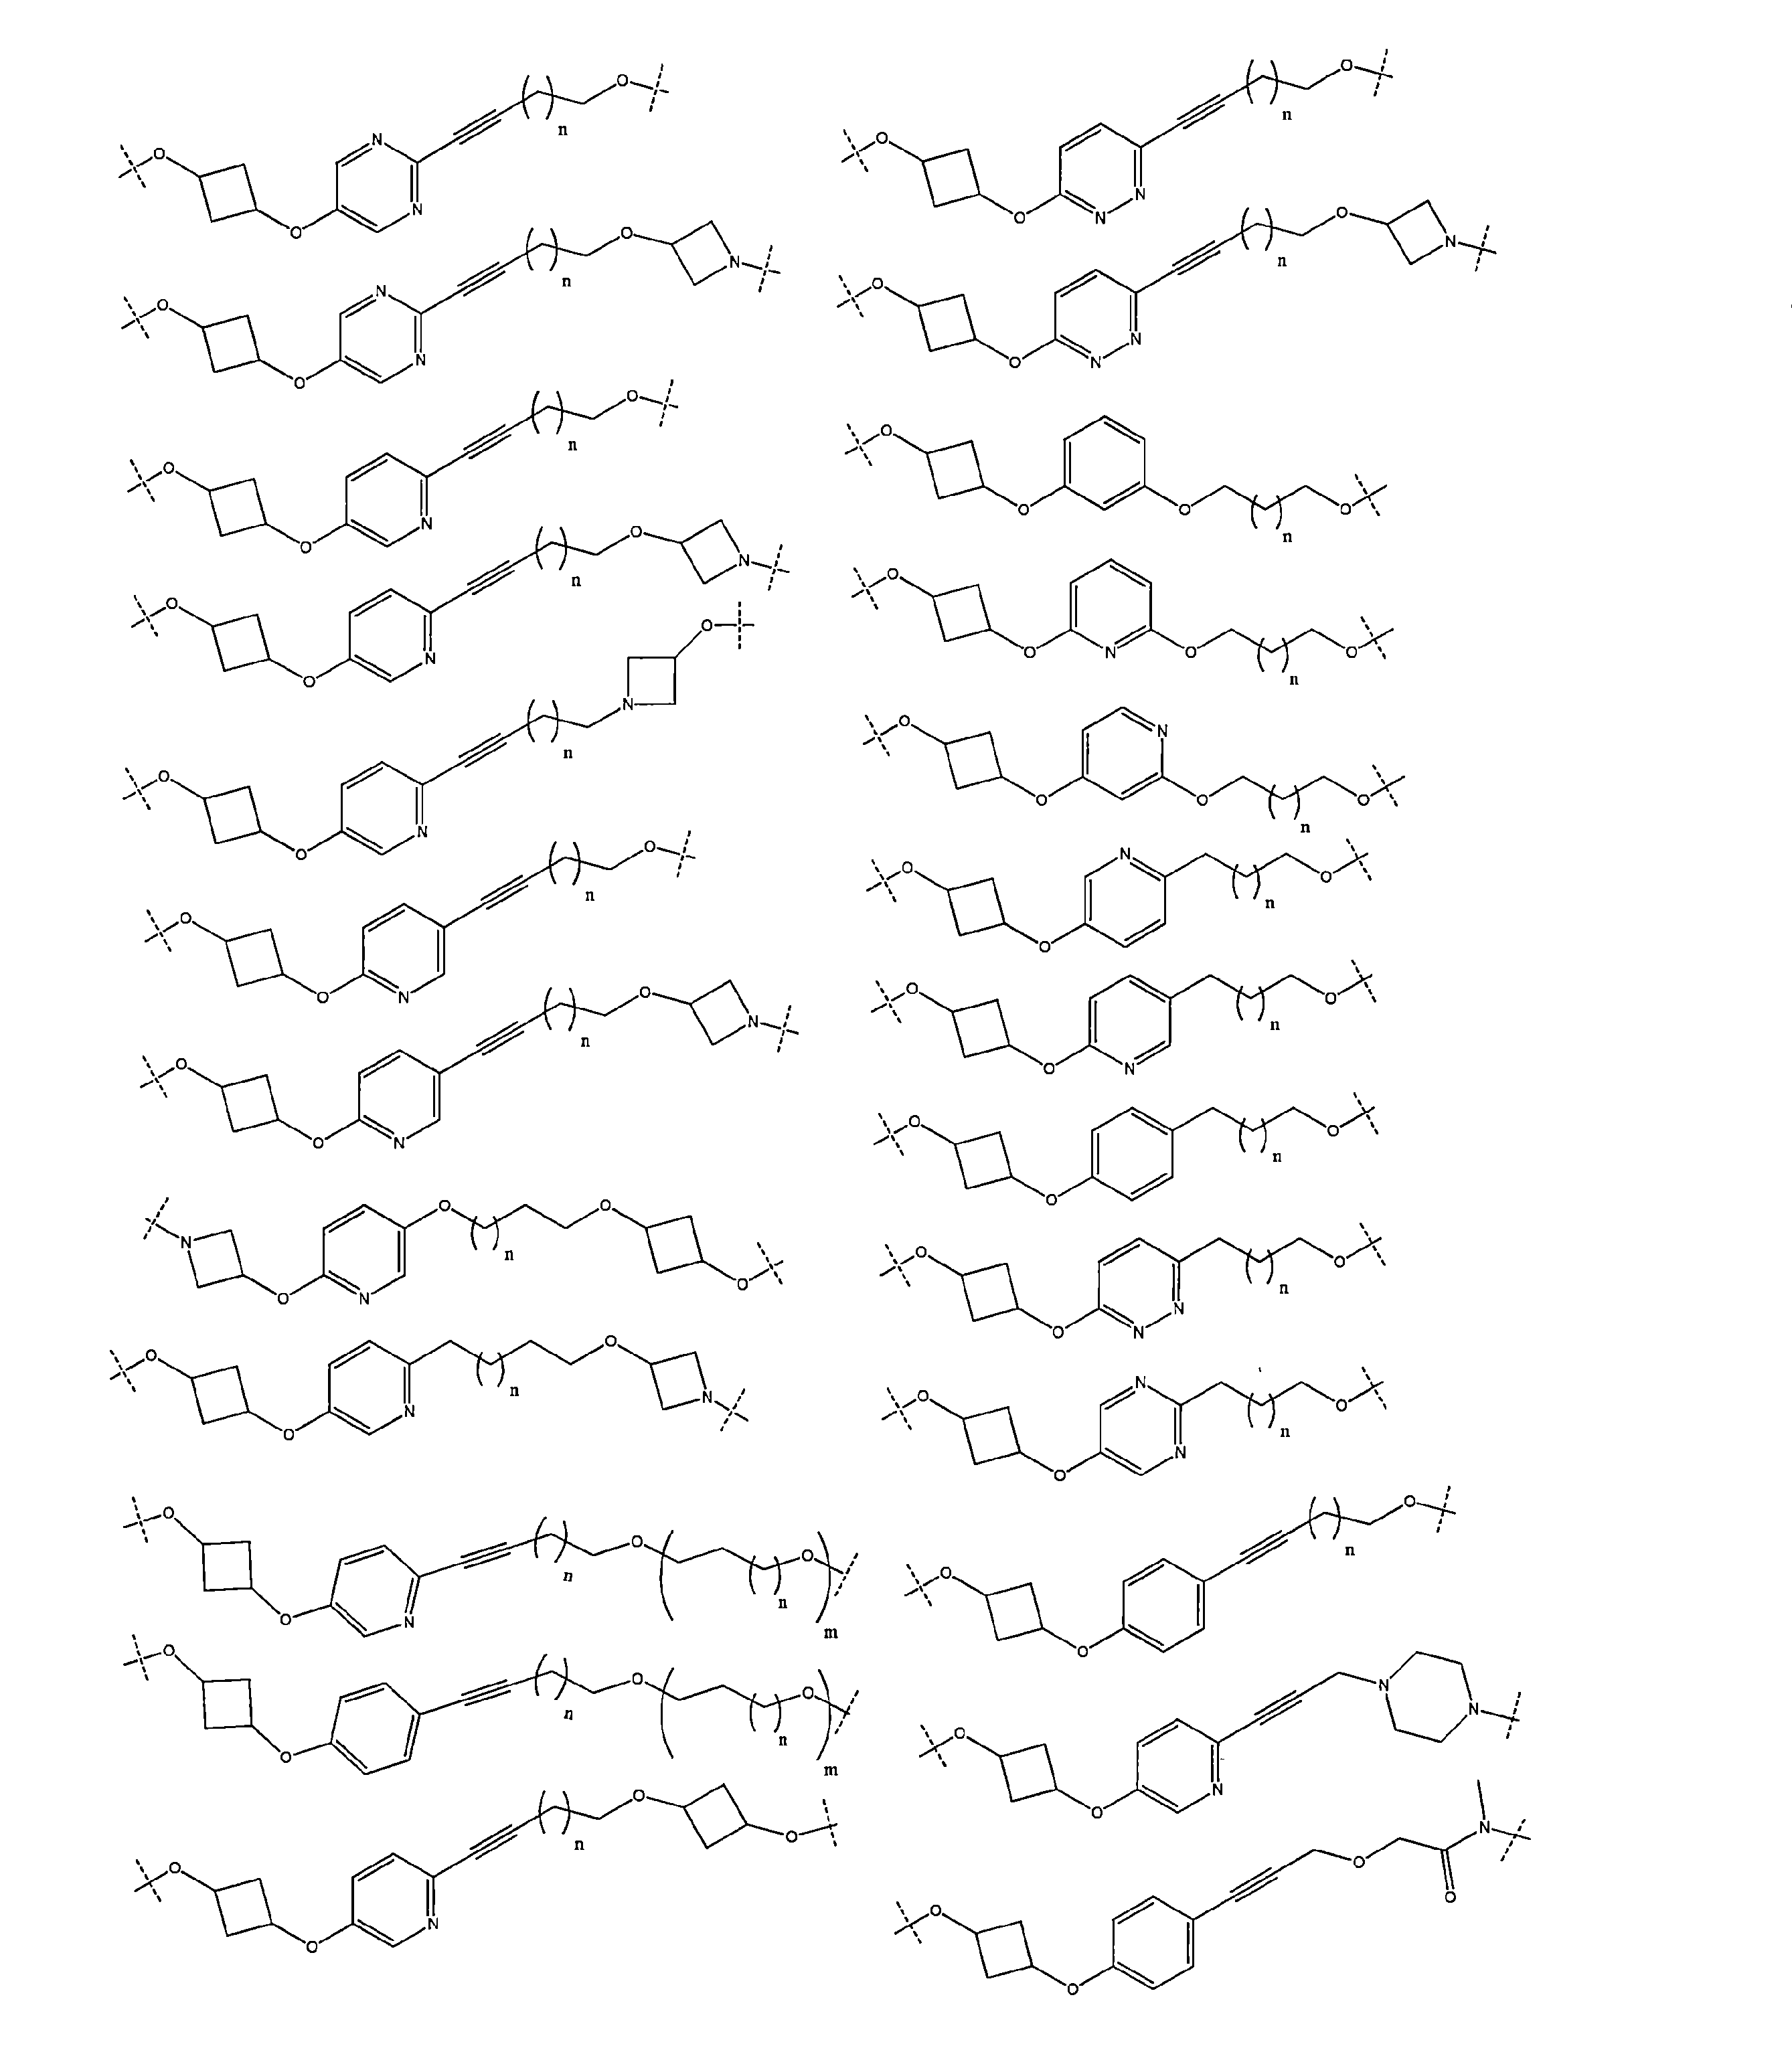


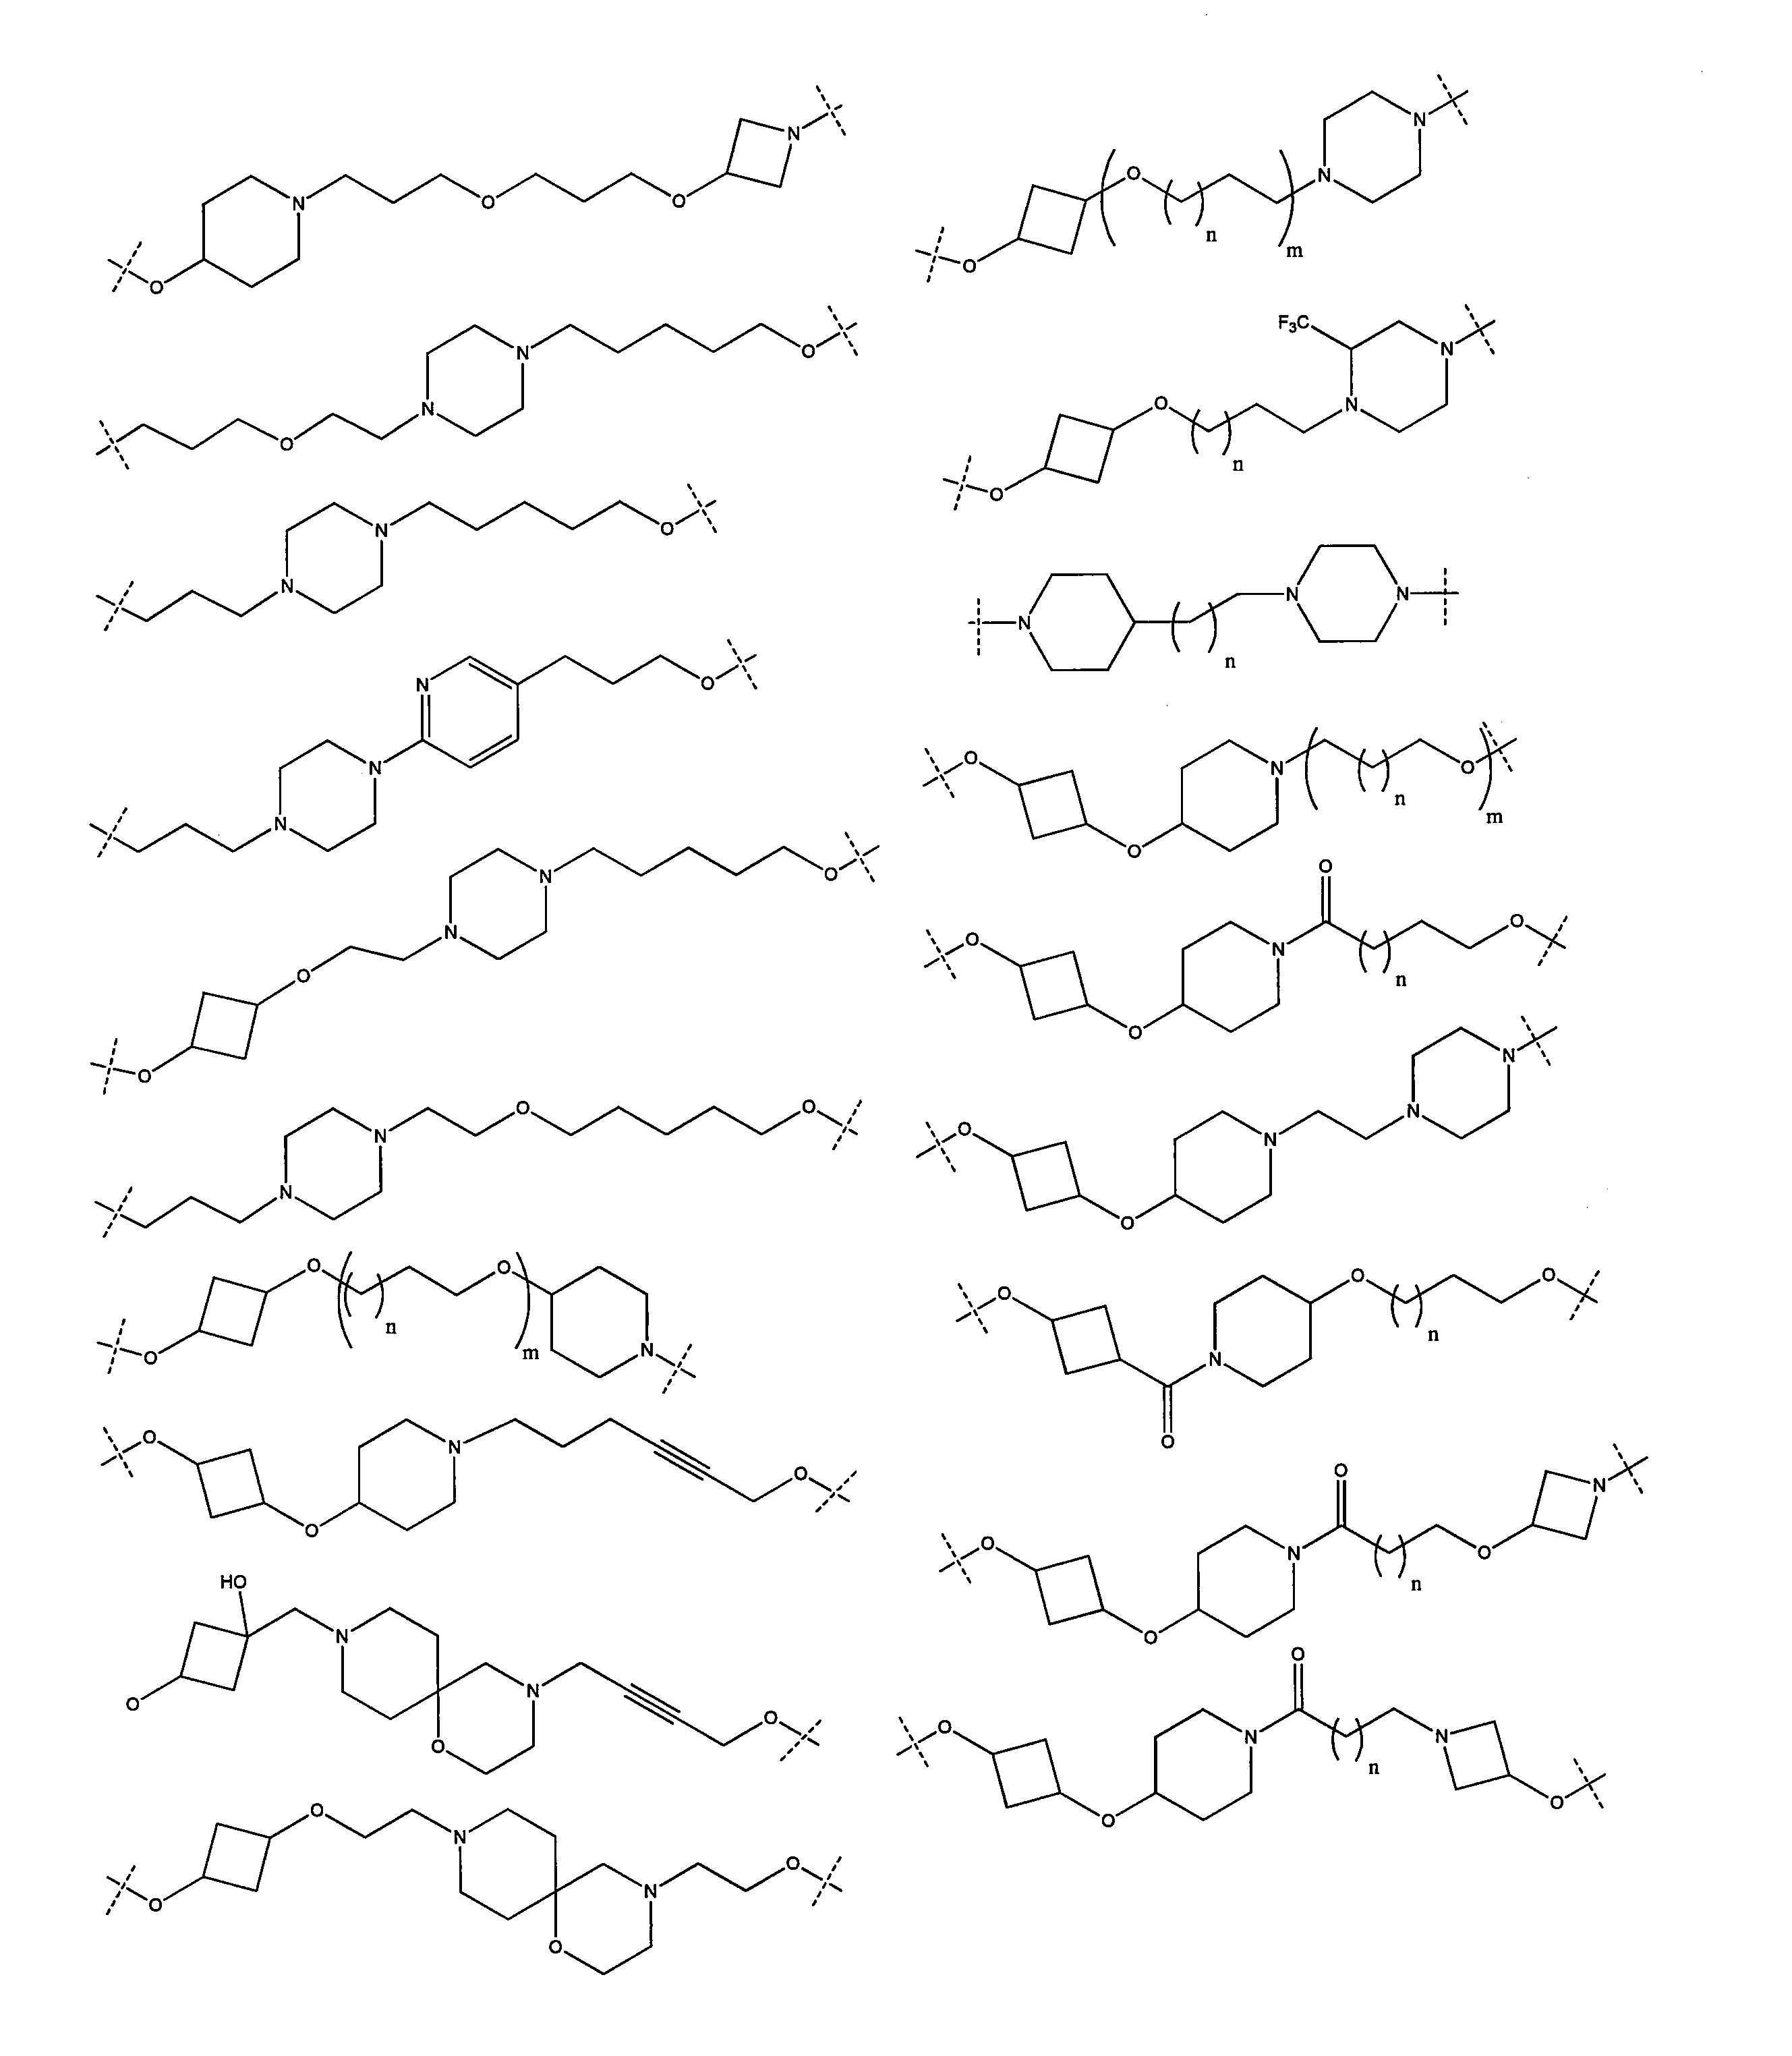


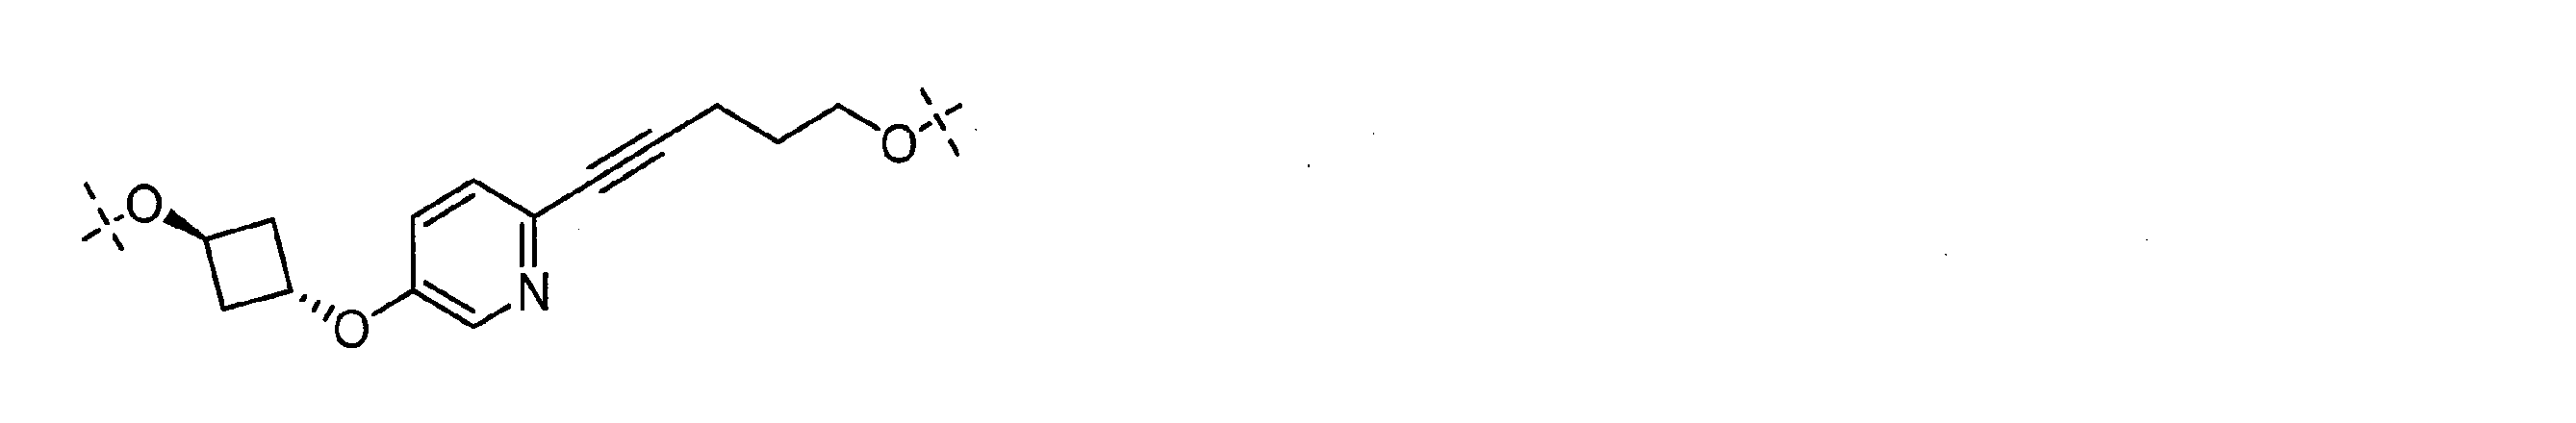
本明細書に記載される任意の態様または実施形態では、リンカー(L)の単位ALは、以下からなる群から選択され、
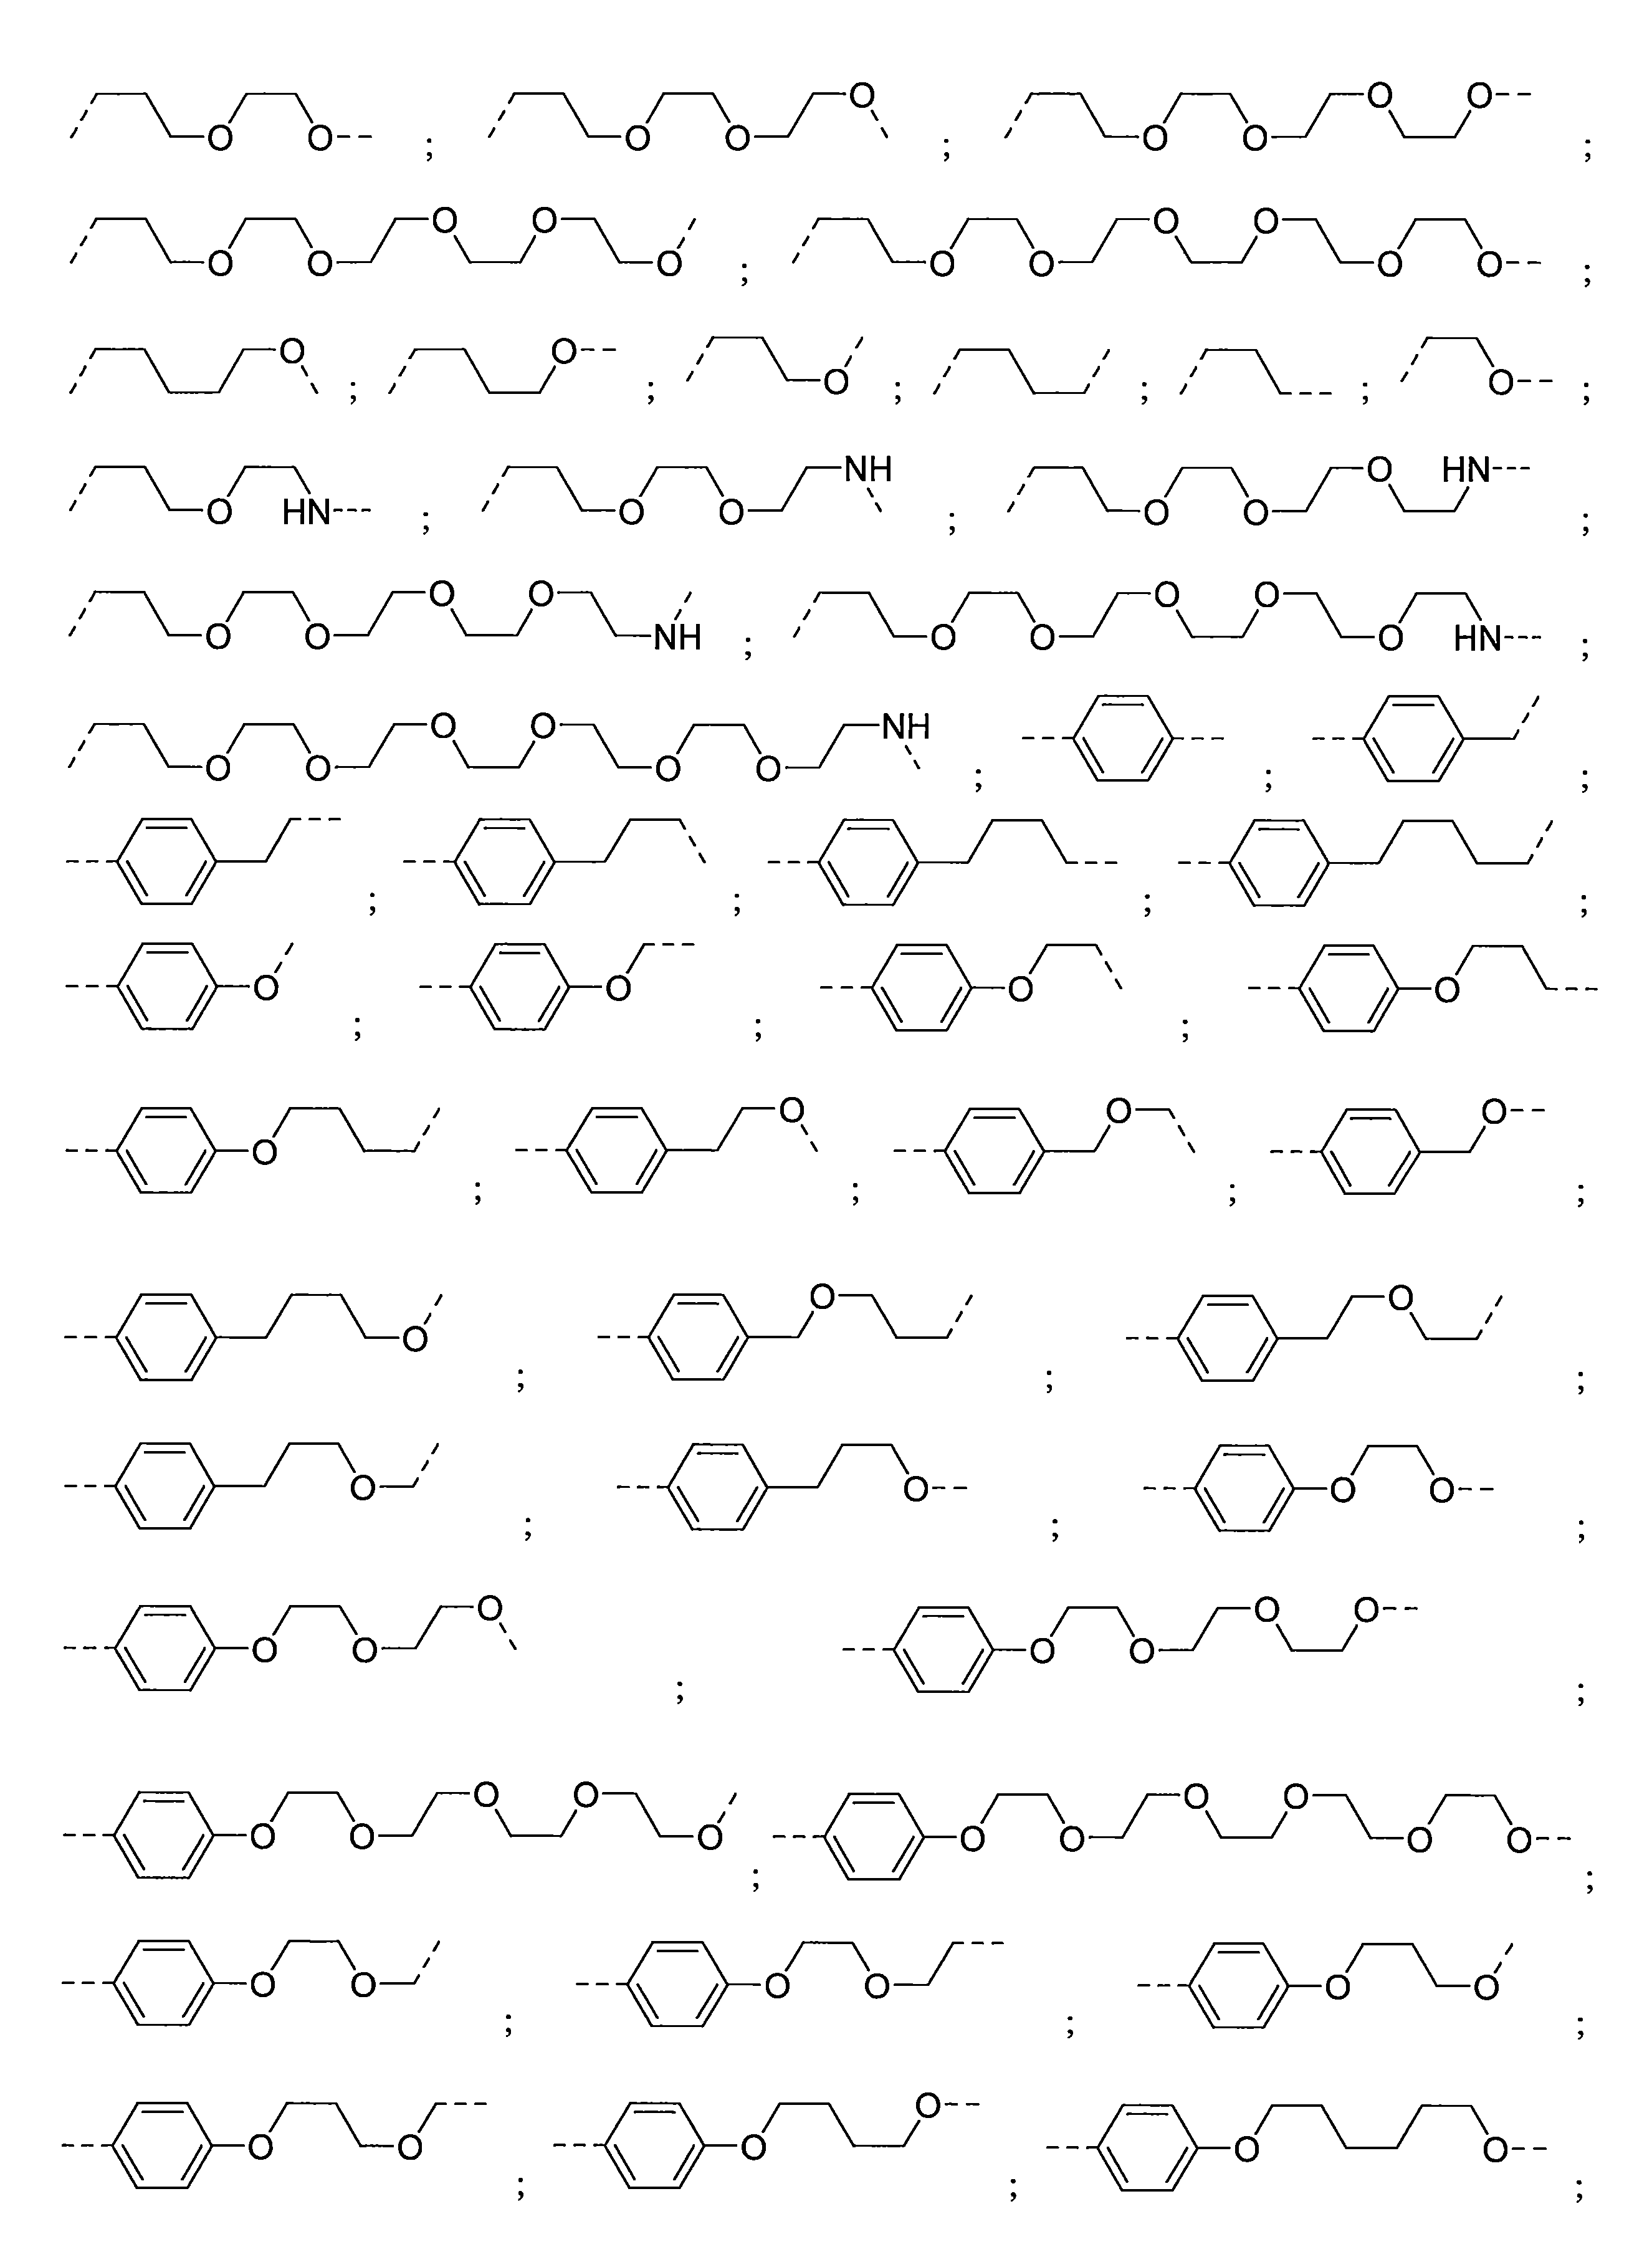

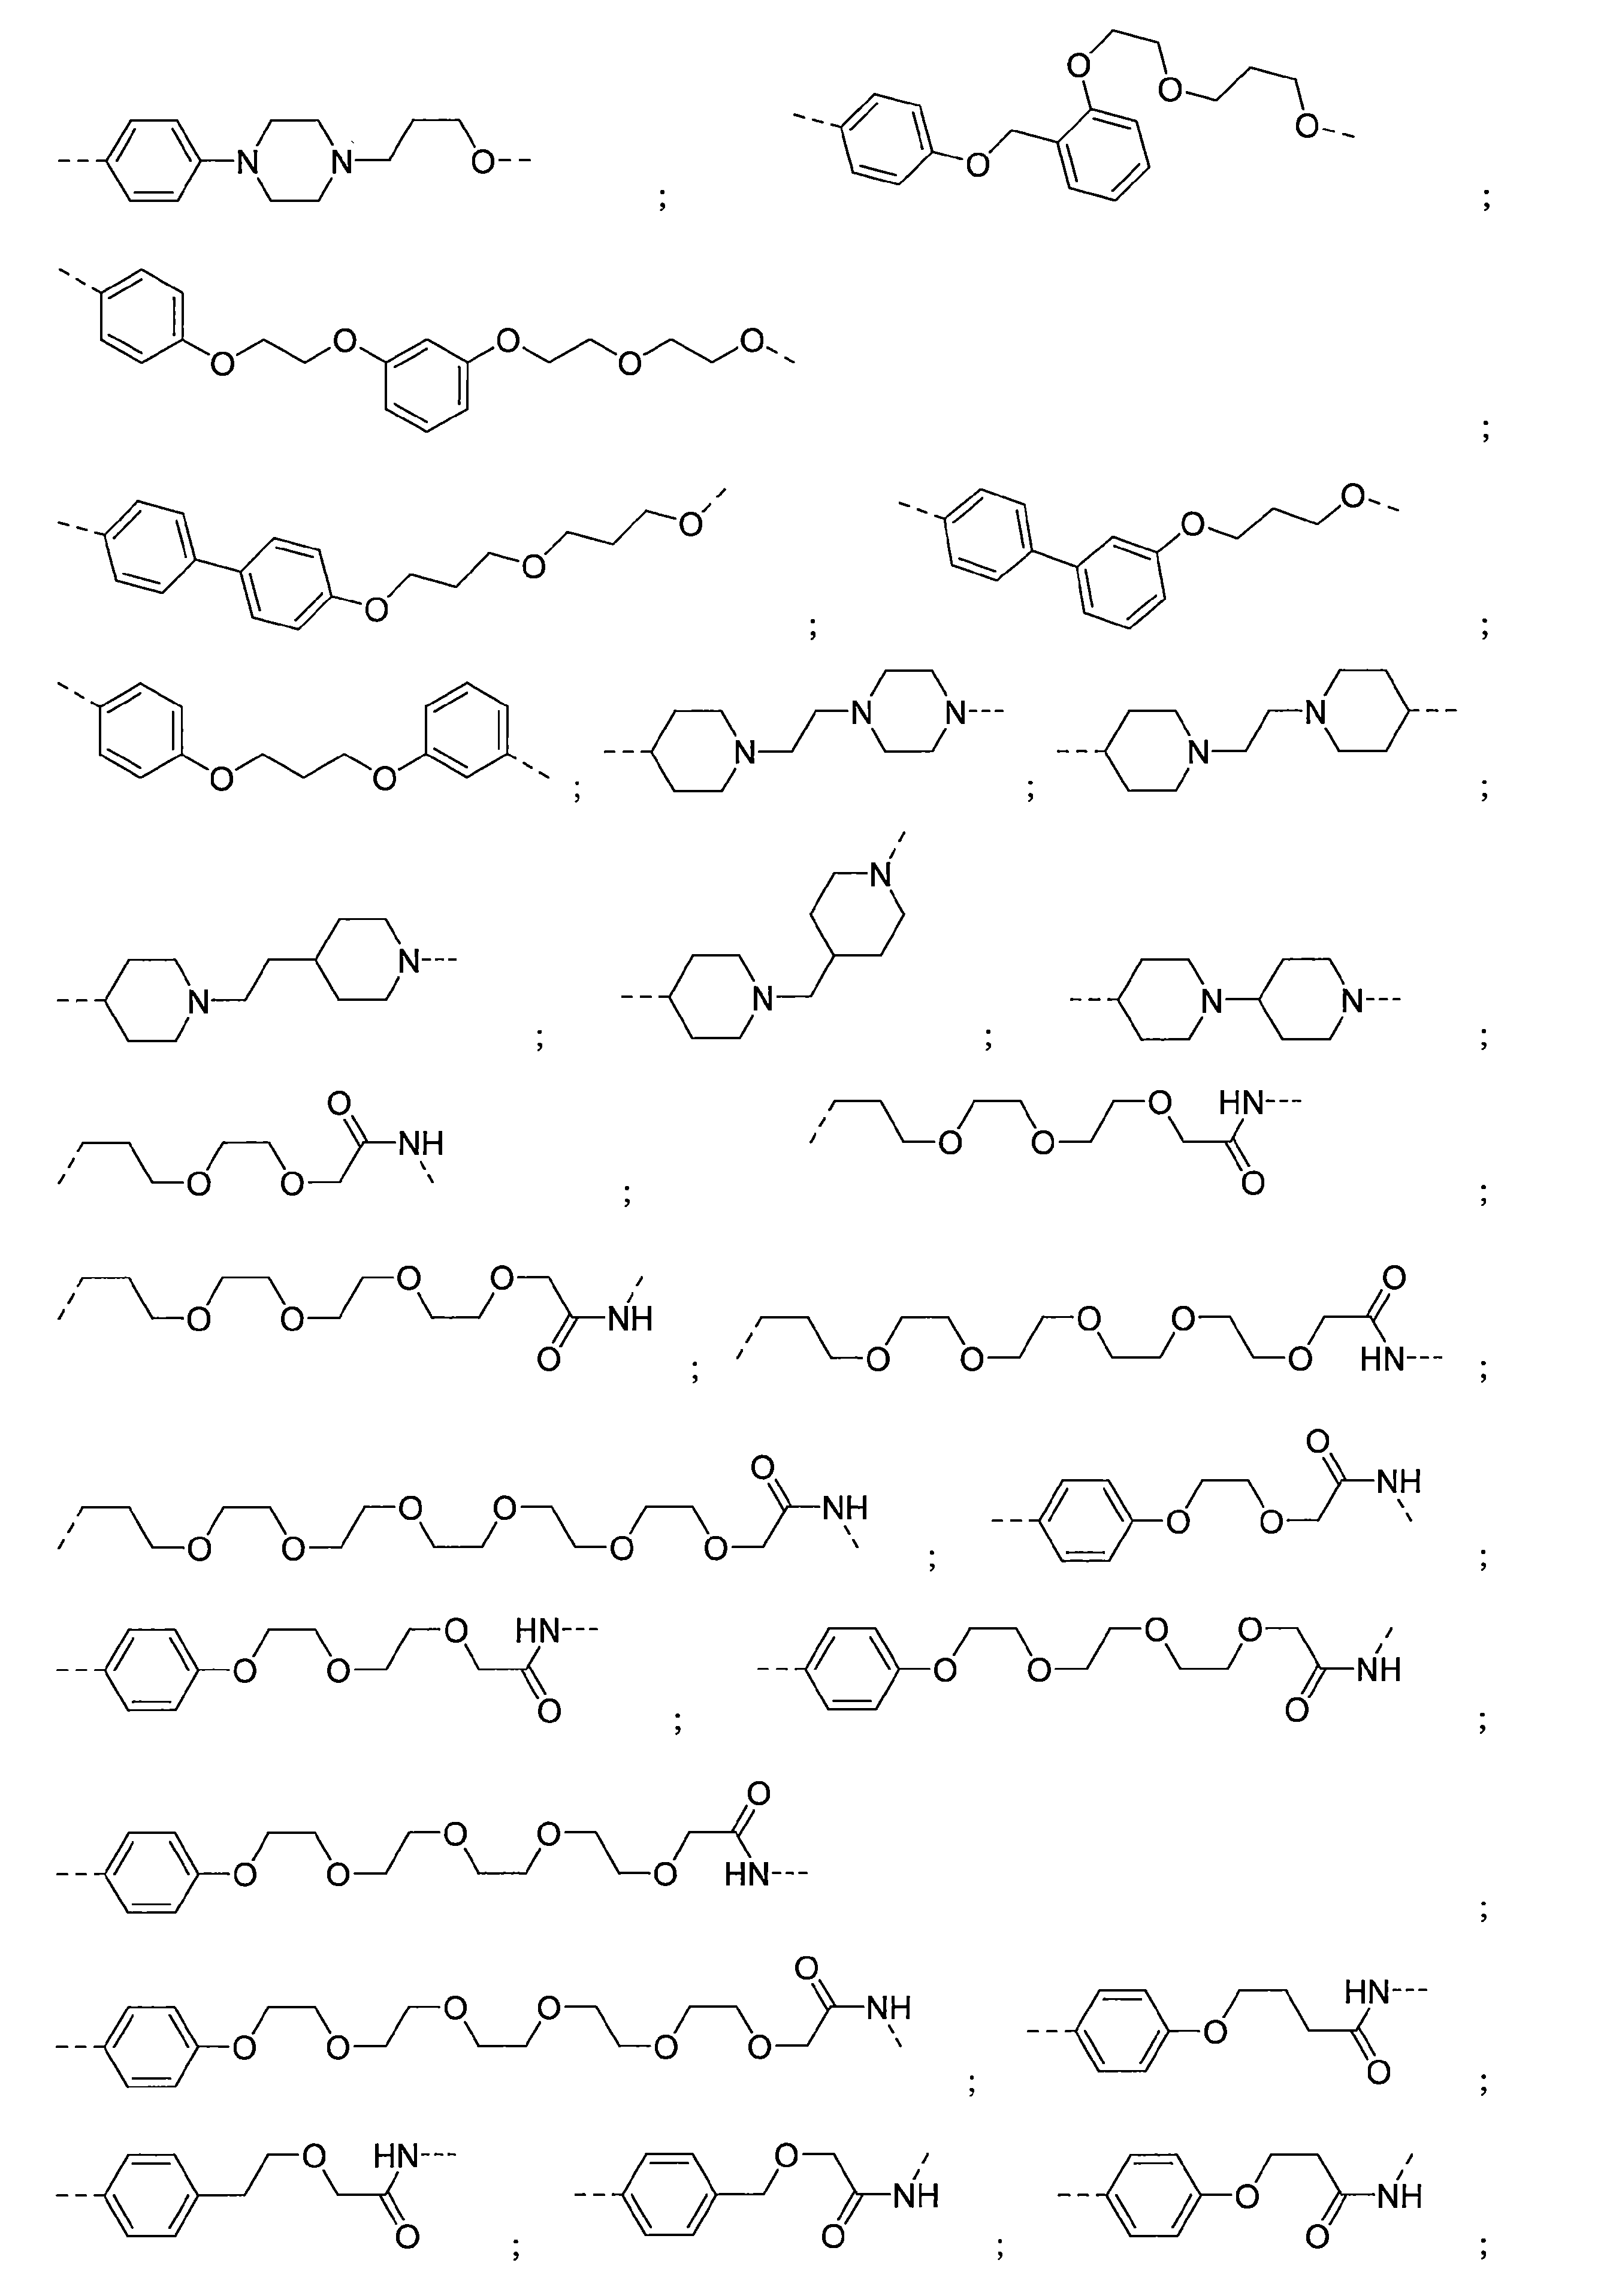
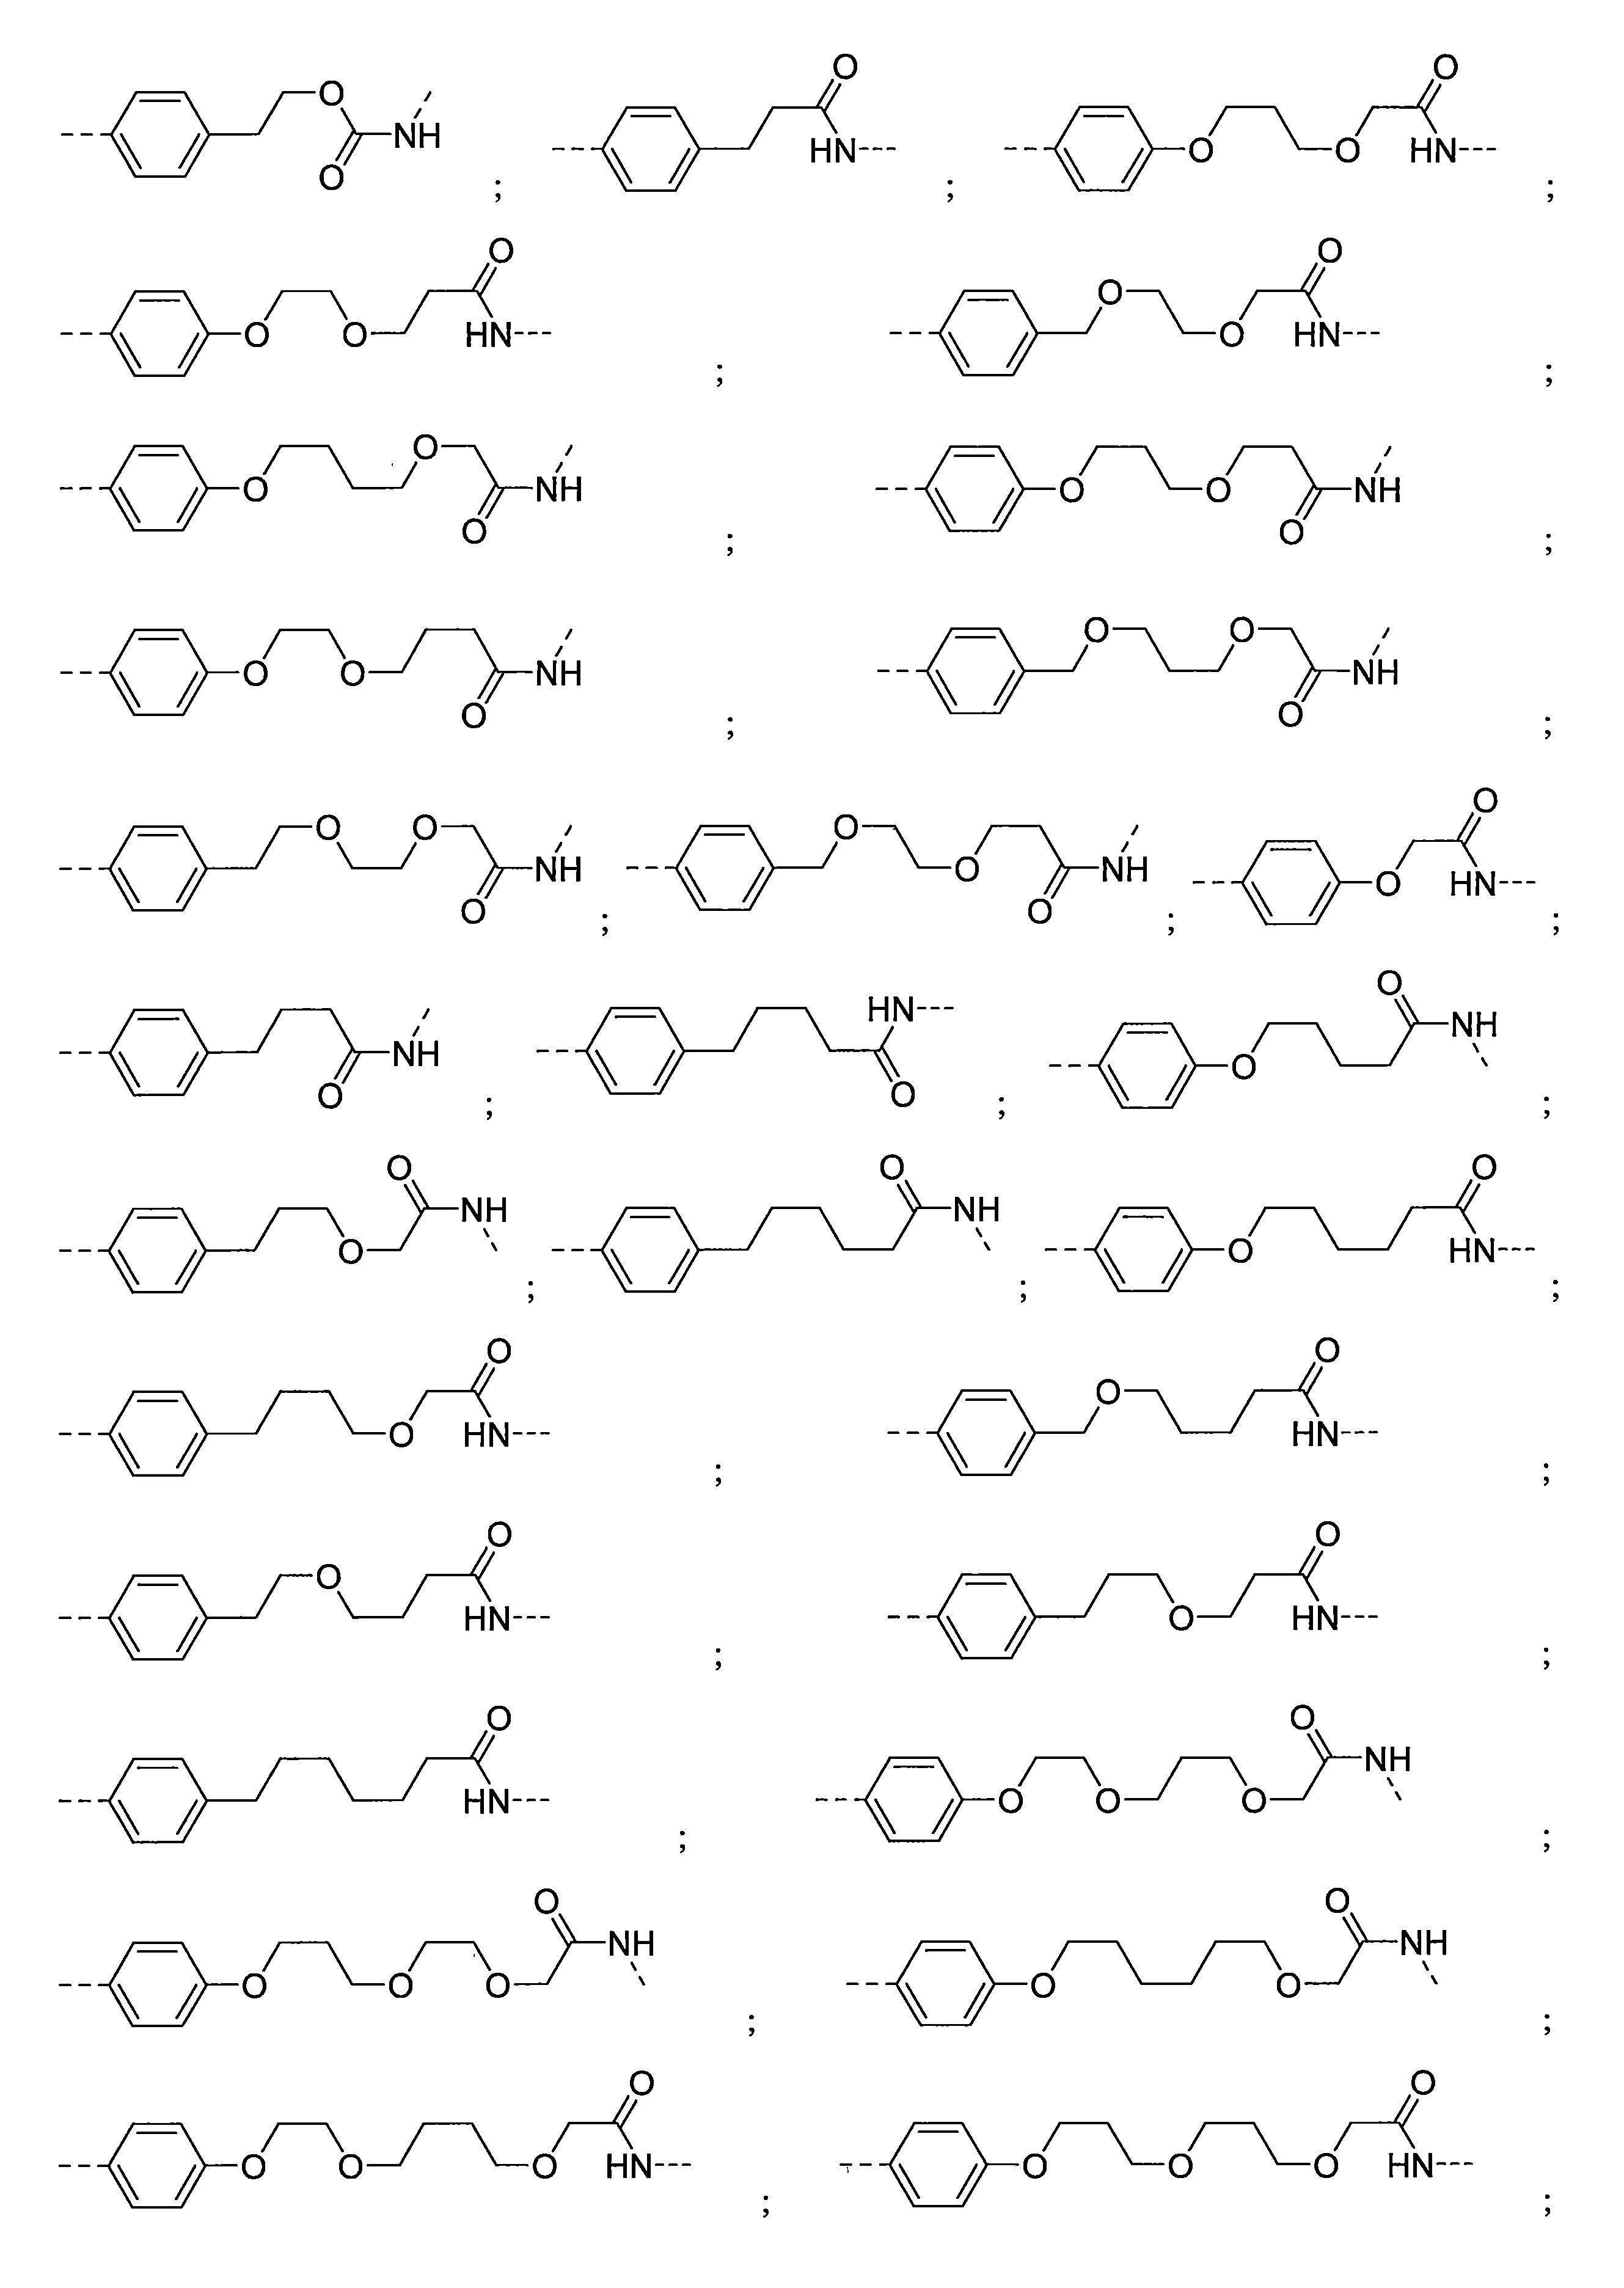
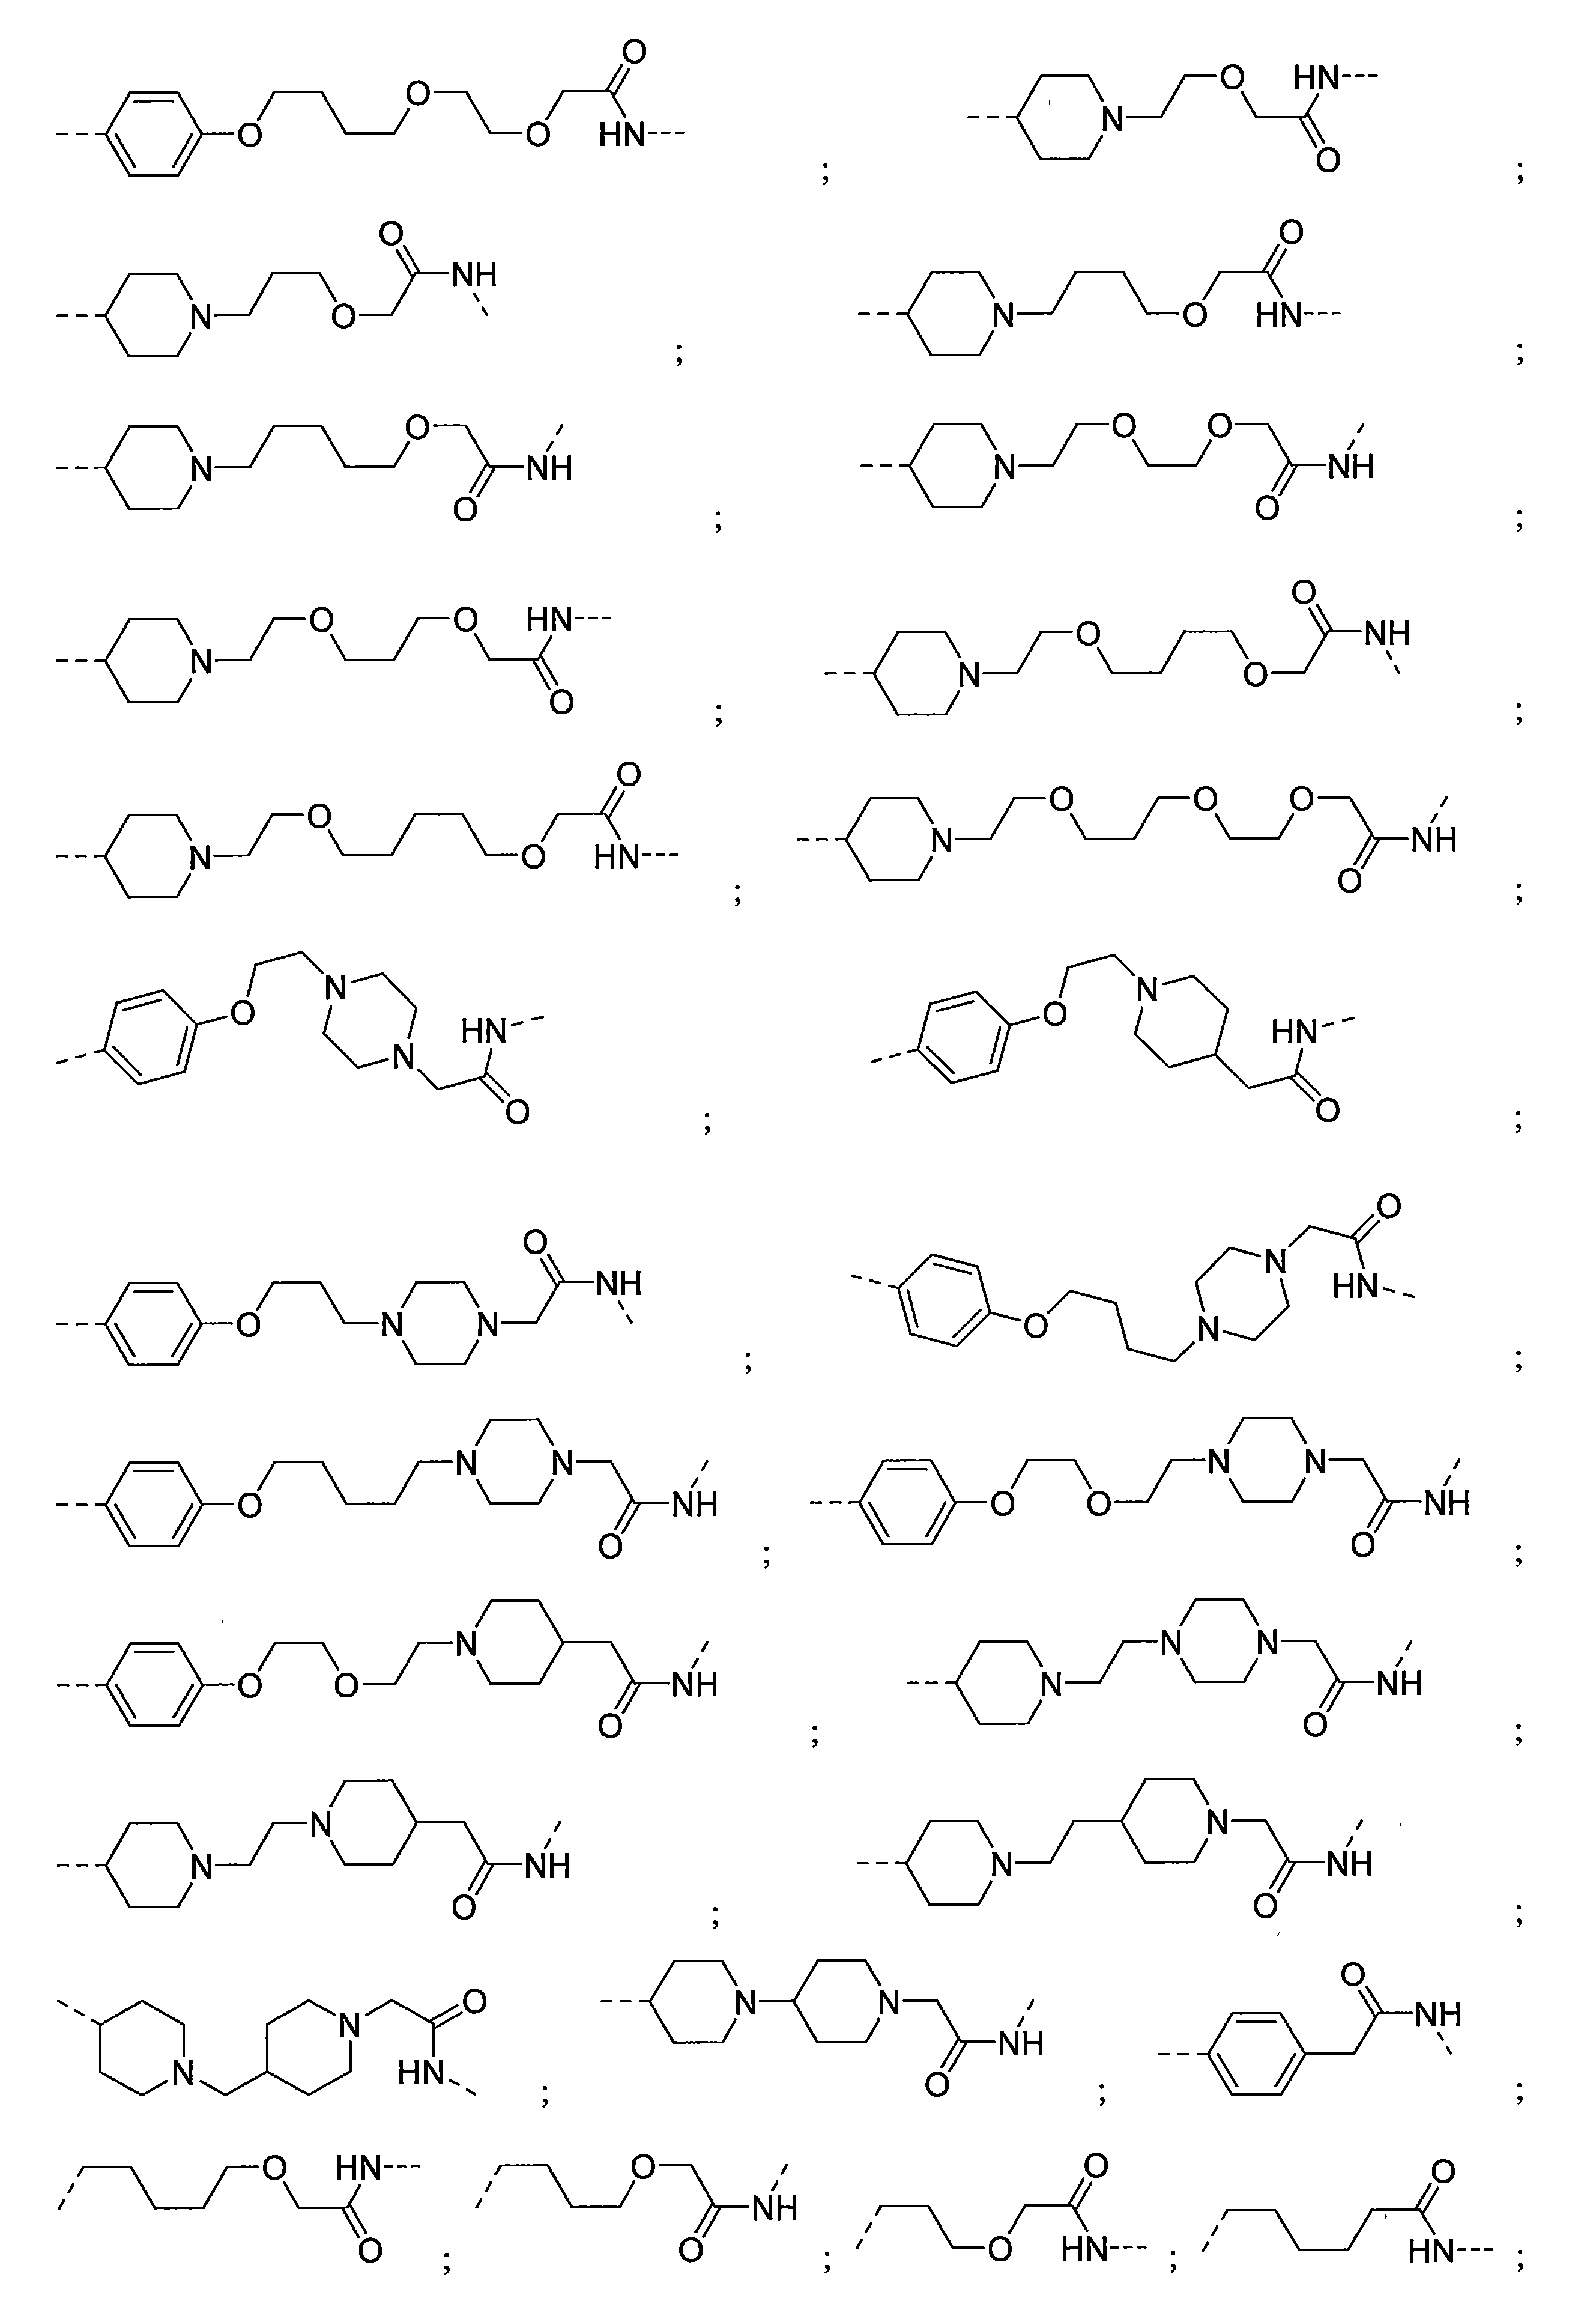


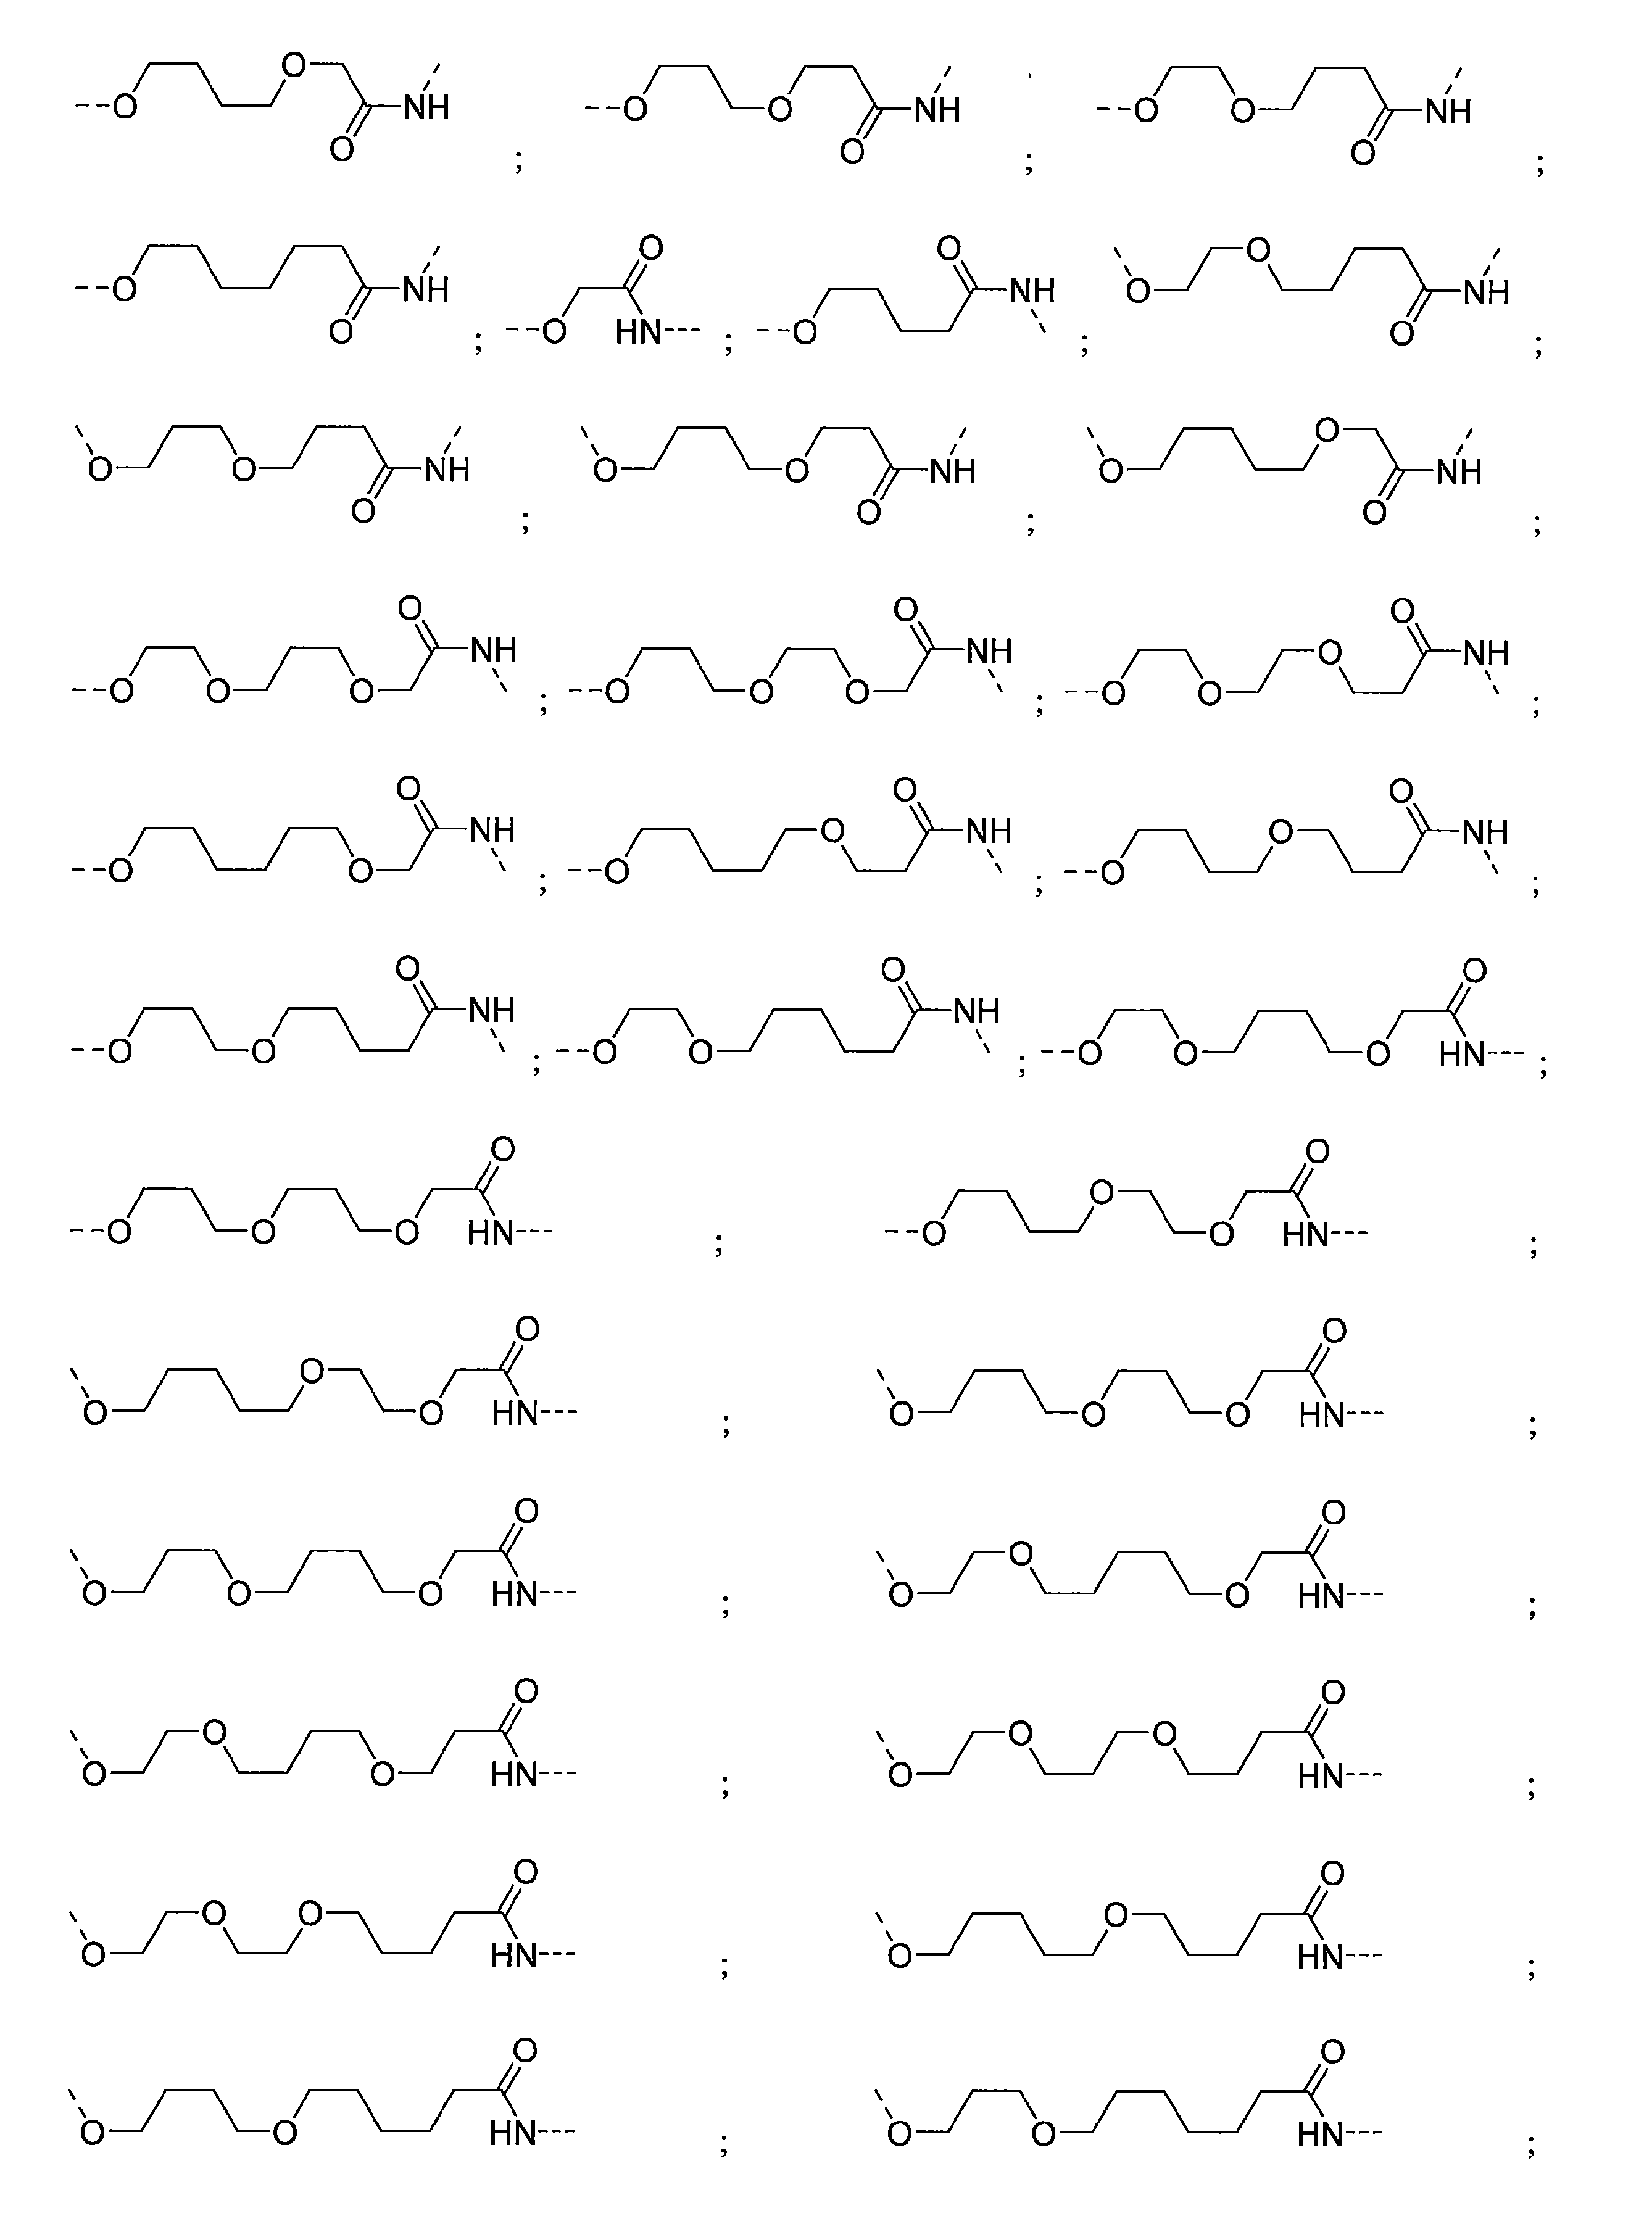


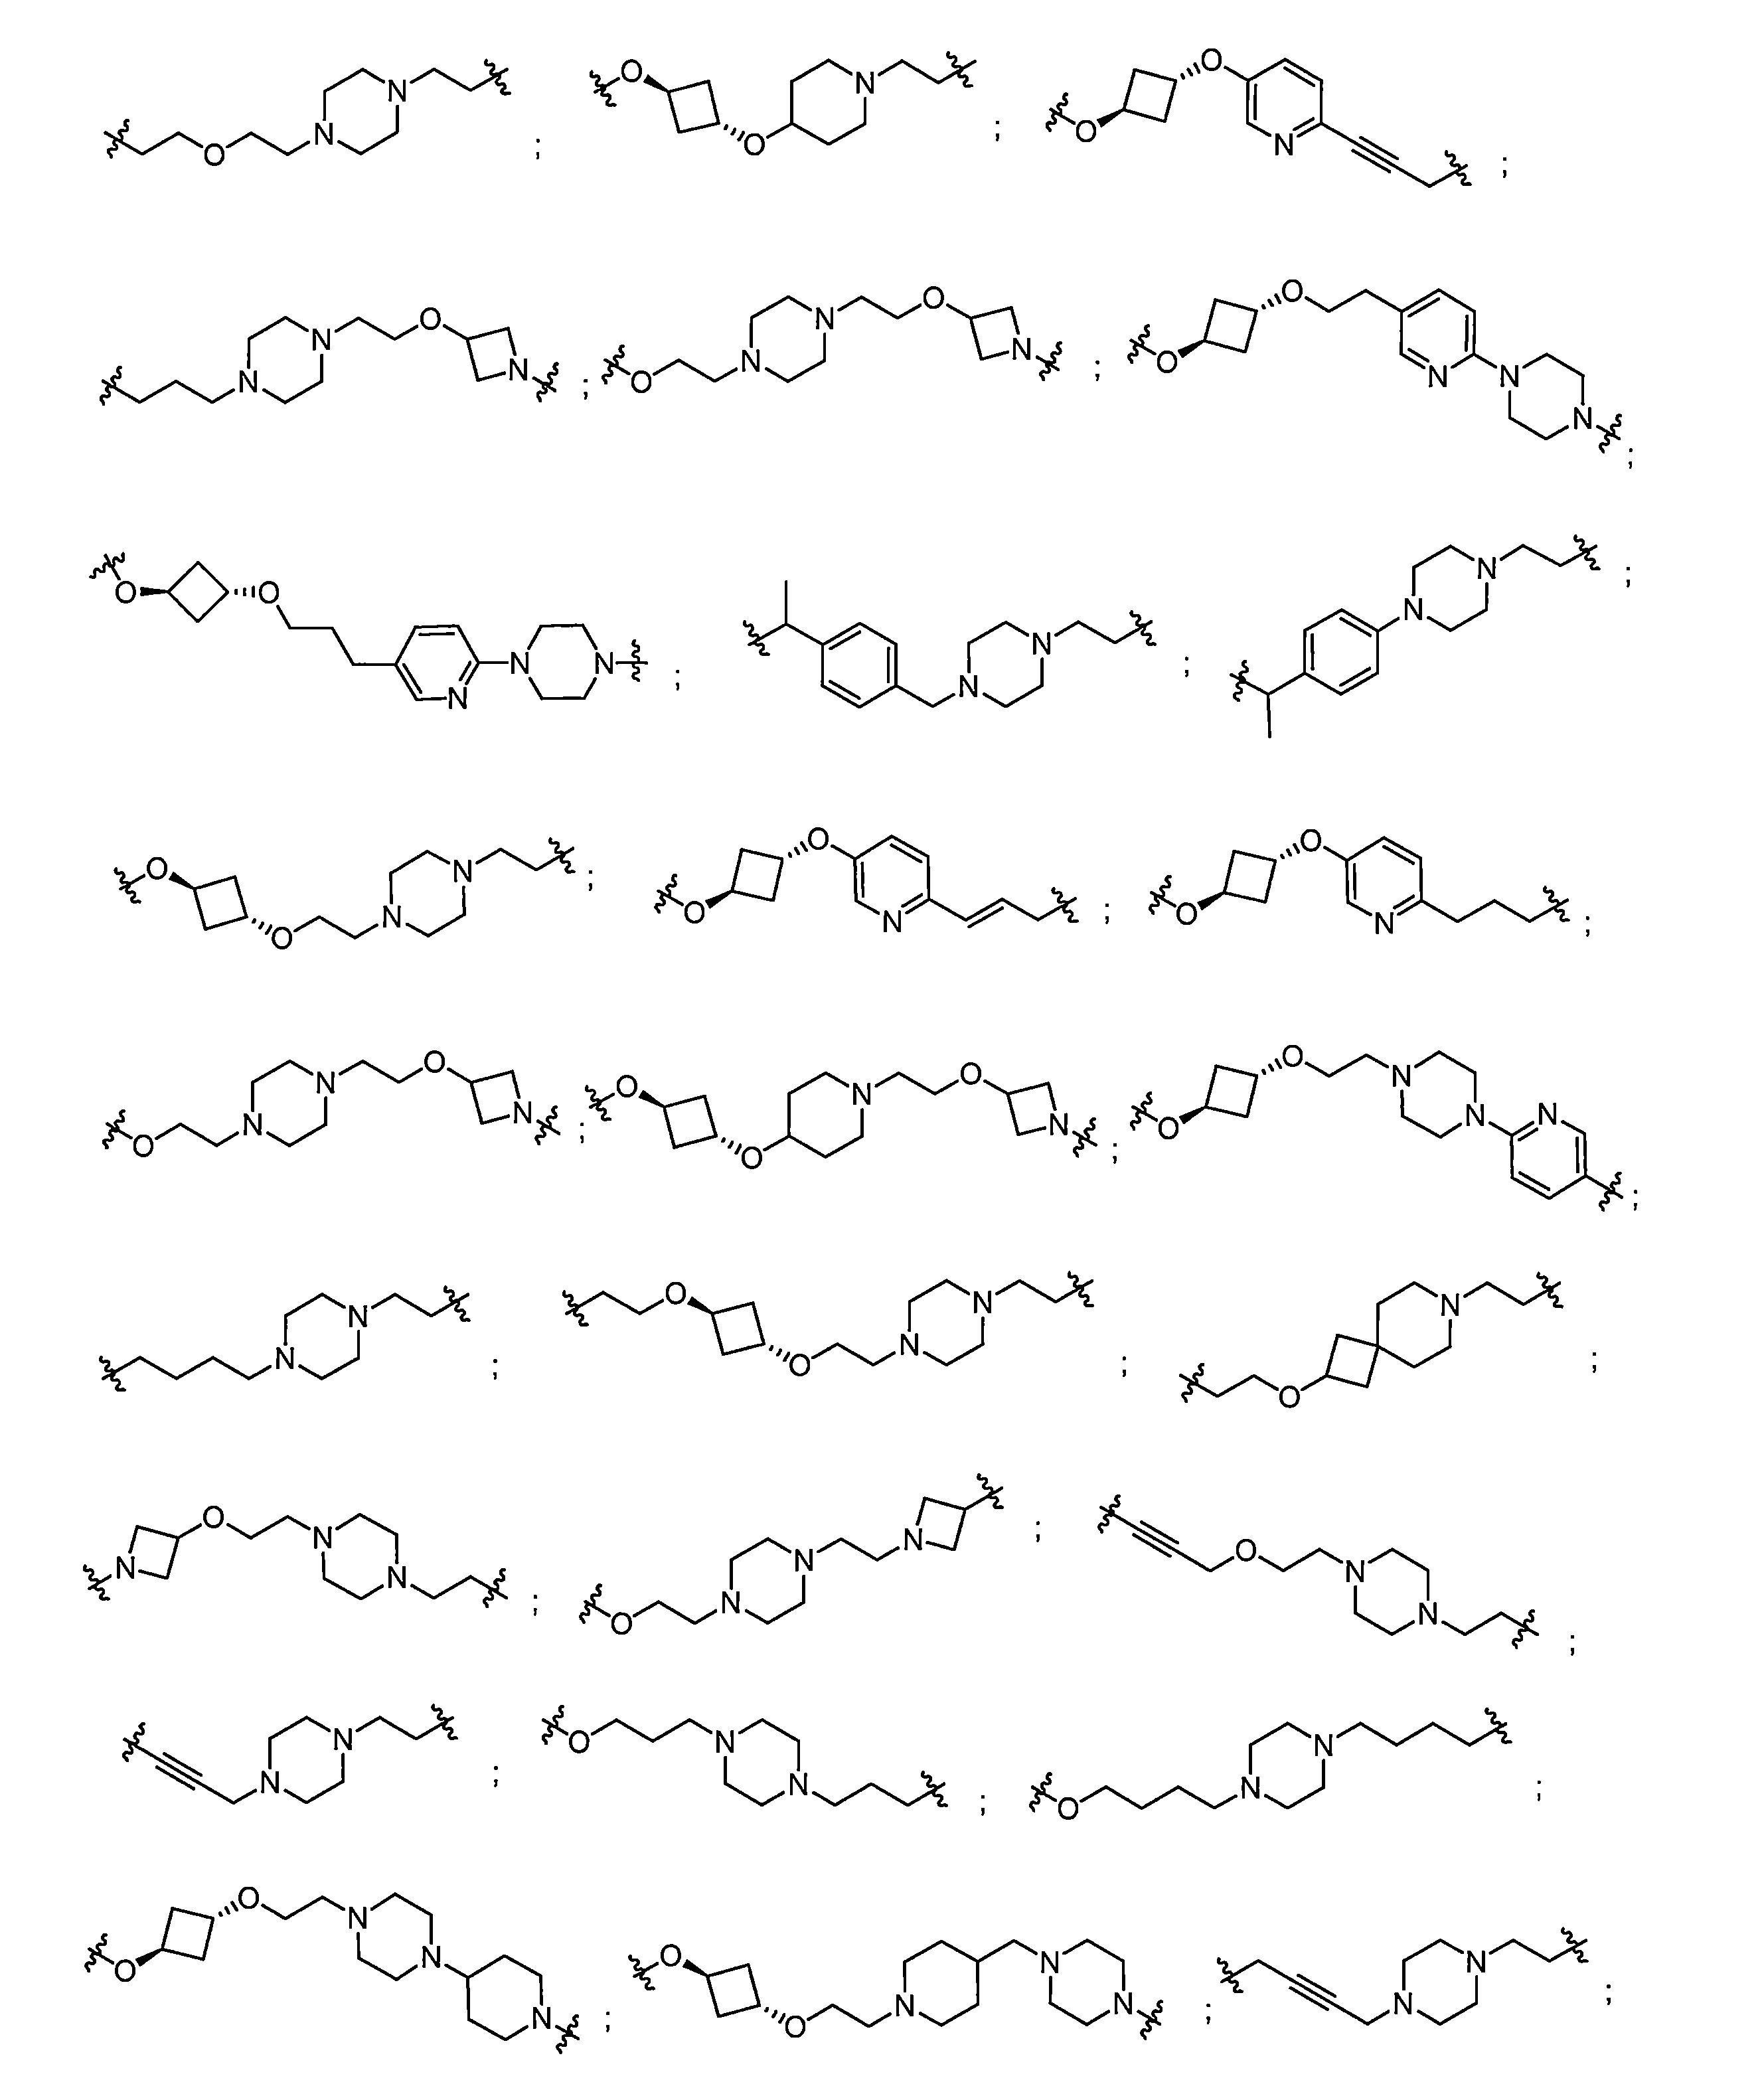


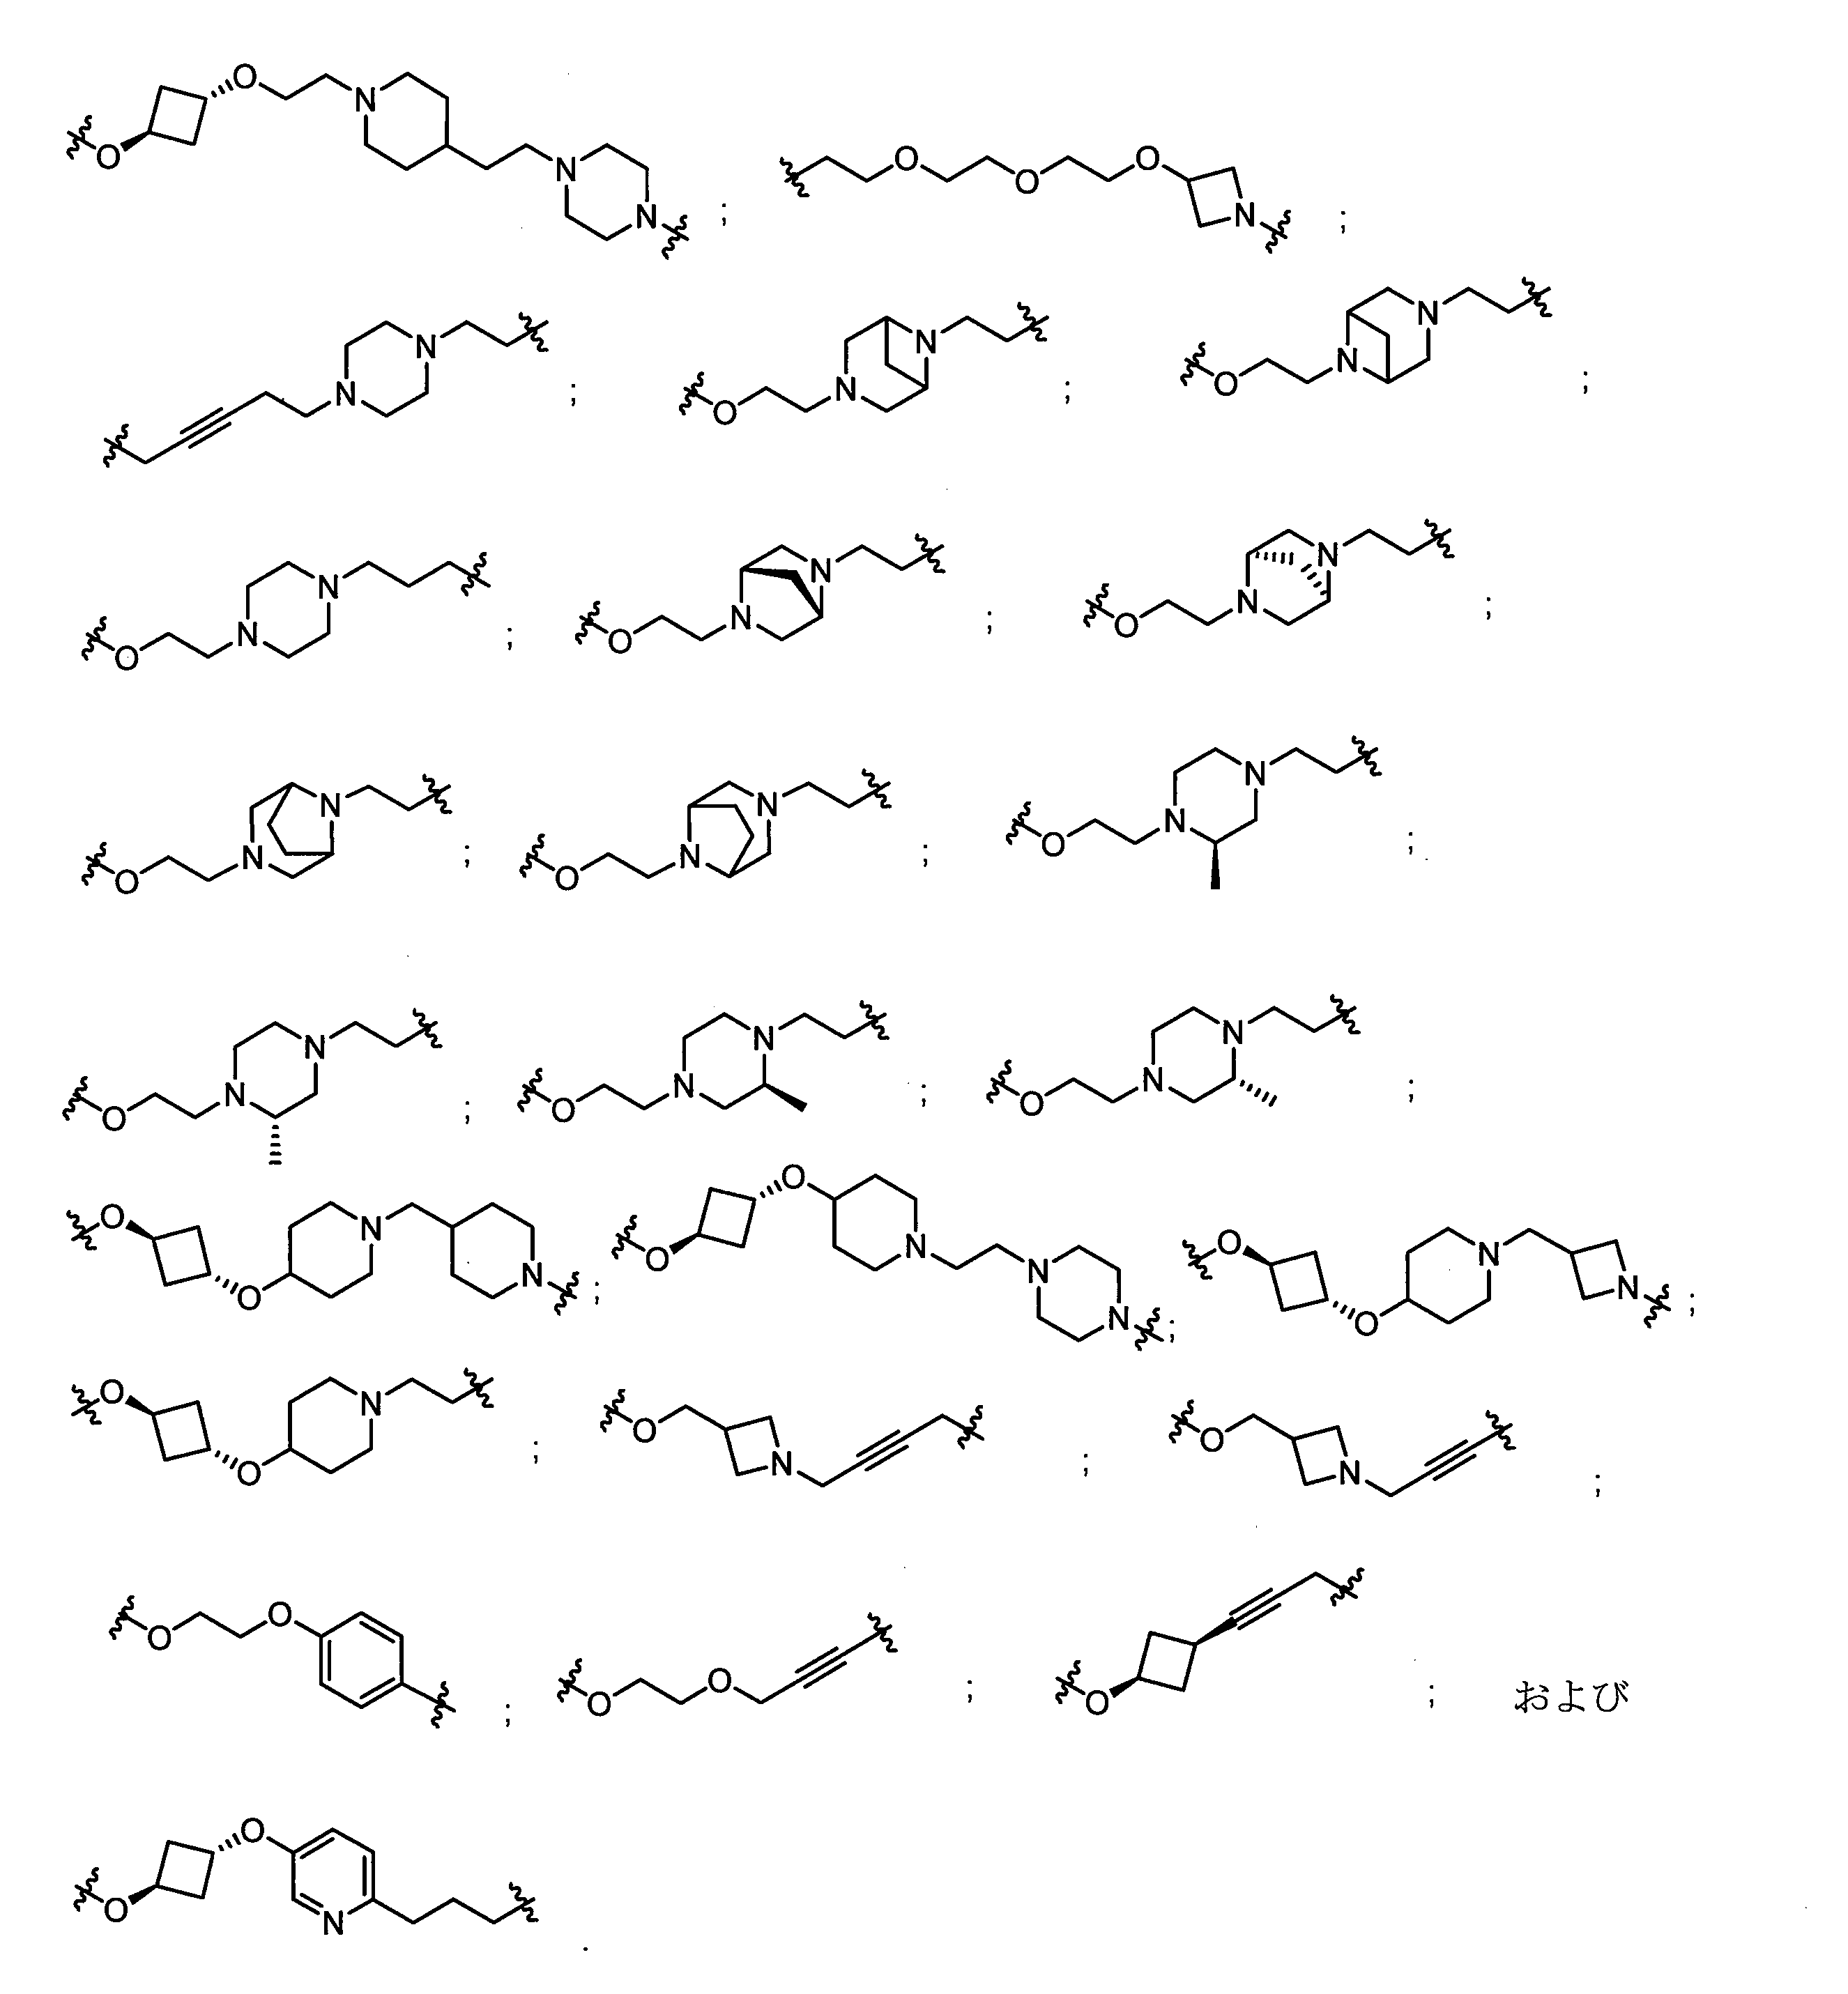
本明細書に記載される任意の態様または実施形態では、リンカー単位またはリンカー(L)は、以下からなる群から選択される構造によって表される基を含み、
-O-(CH2)m-O(CH2)n-O(CH2)o-O(CH2)p-O(CH2)q-O(CH2)r-O(CH2)s-O(CH2)t-、
-O-(CH2)m-O(CH2)n-O(CH2)o-O(CH2)p-O(CH2)q-O(CH2)r-O(CH2)s-O-、
-(CH2)m-O(CH2)n-O(CH2)o-O(CH2)p-O(CH2)q-O(CH2)r-O(CH2)s-O(CH2)t-、
-CH=CH(CH2)m-O(CH2)n-O(CH2)o-O(CH2)p-O(CH2)q-O(CH2)r-O(CH2)s-O(CH2)t-、


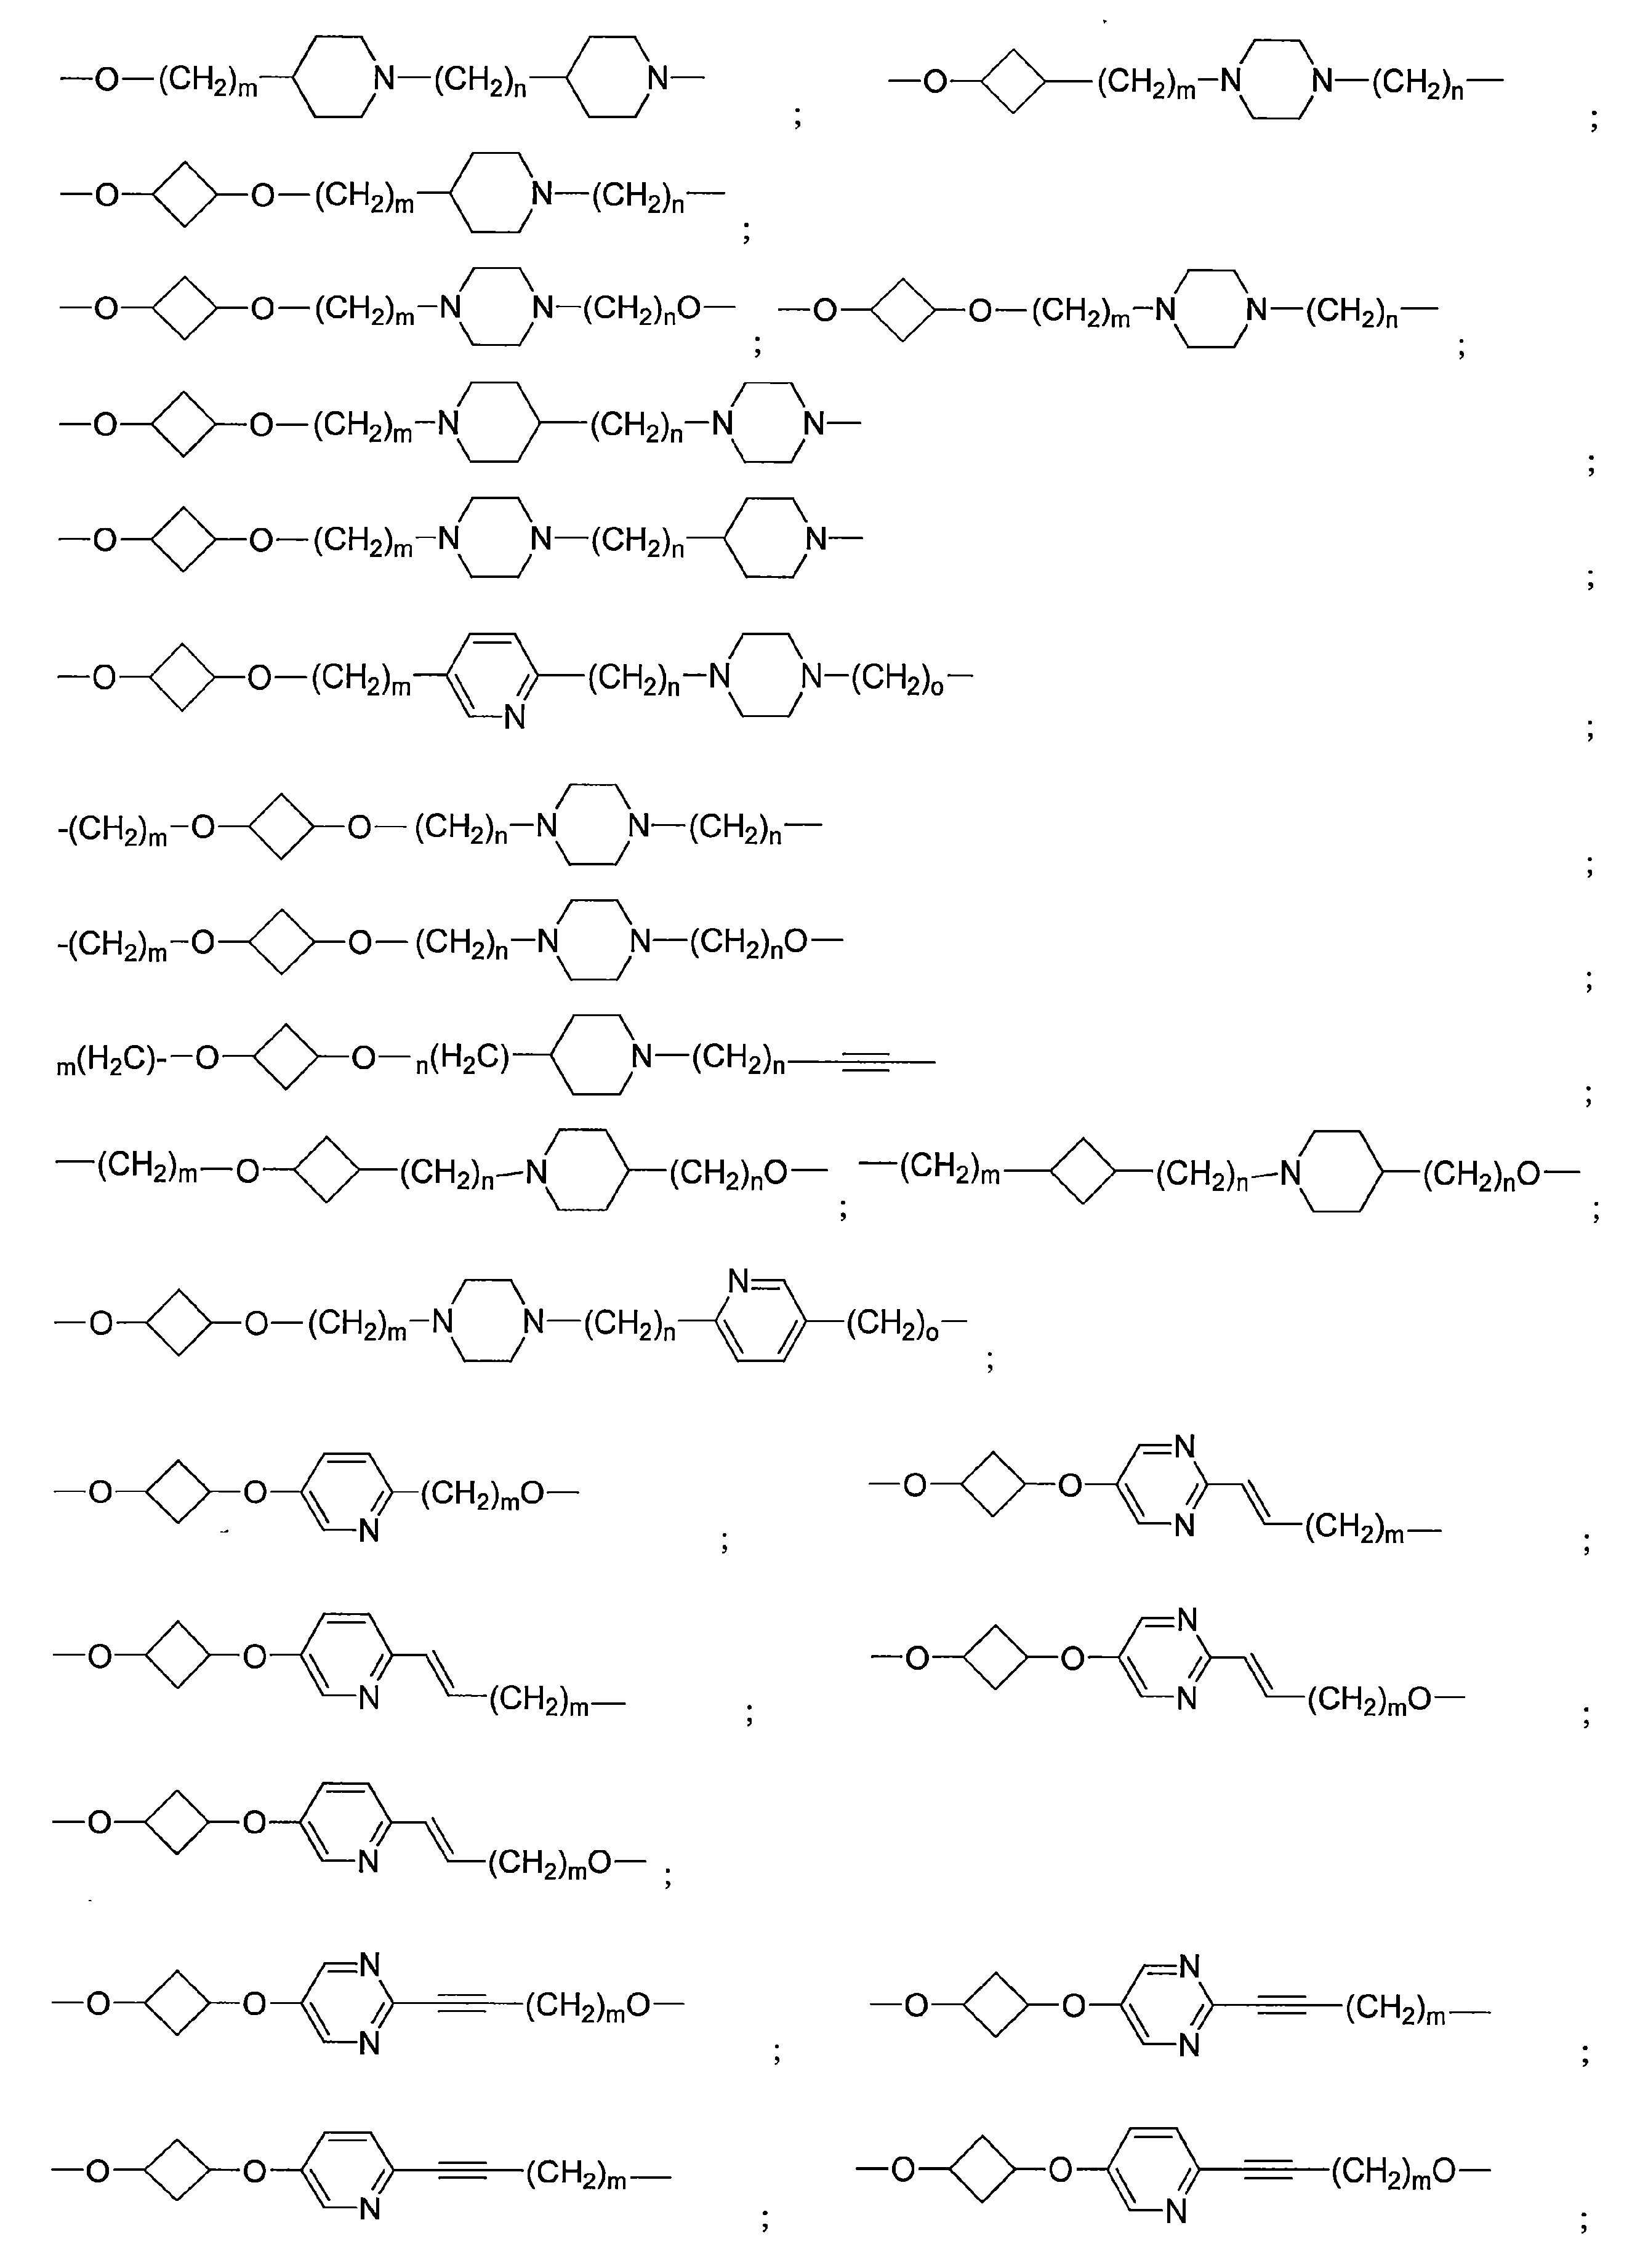


-O-(CH2)m-O(CH2)n-O(CH2)o-O(CH2)p-O(CH2)q-O(CH2)r-O(CH2)s-O(CH2)t-,
—O—(CH2)m—O(CH2)n—O(CH2)o—O(CH2)p—O(CH2)q—O(CH2)r—O(CH2)s—O—,
-(CH2)m-O(CH2)n-O(CH2)o-O(CH2)p-O(CH2)q-O(CH2)r-O(CH2)s-O(CH2)t-,
-CH=CH(CH2)m-O(CH2)n-O(CH2)o-O(CH2)p-O(CH2)q-O(CH2)r-O(CH2)s-O(CH2)t-,
本明細書に記載される任意の態様または実施形態において、リンカー(L)は、以下からなる群から選択される:


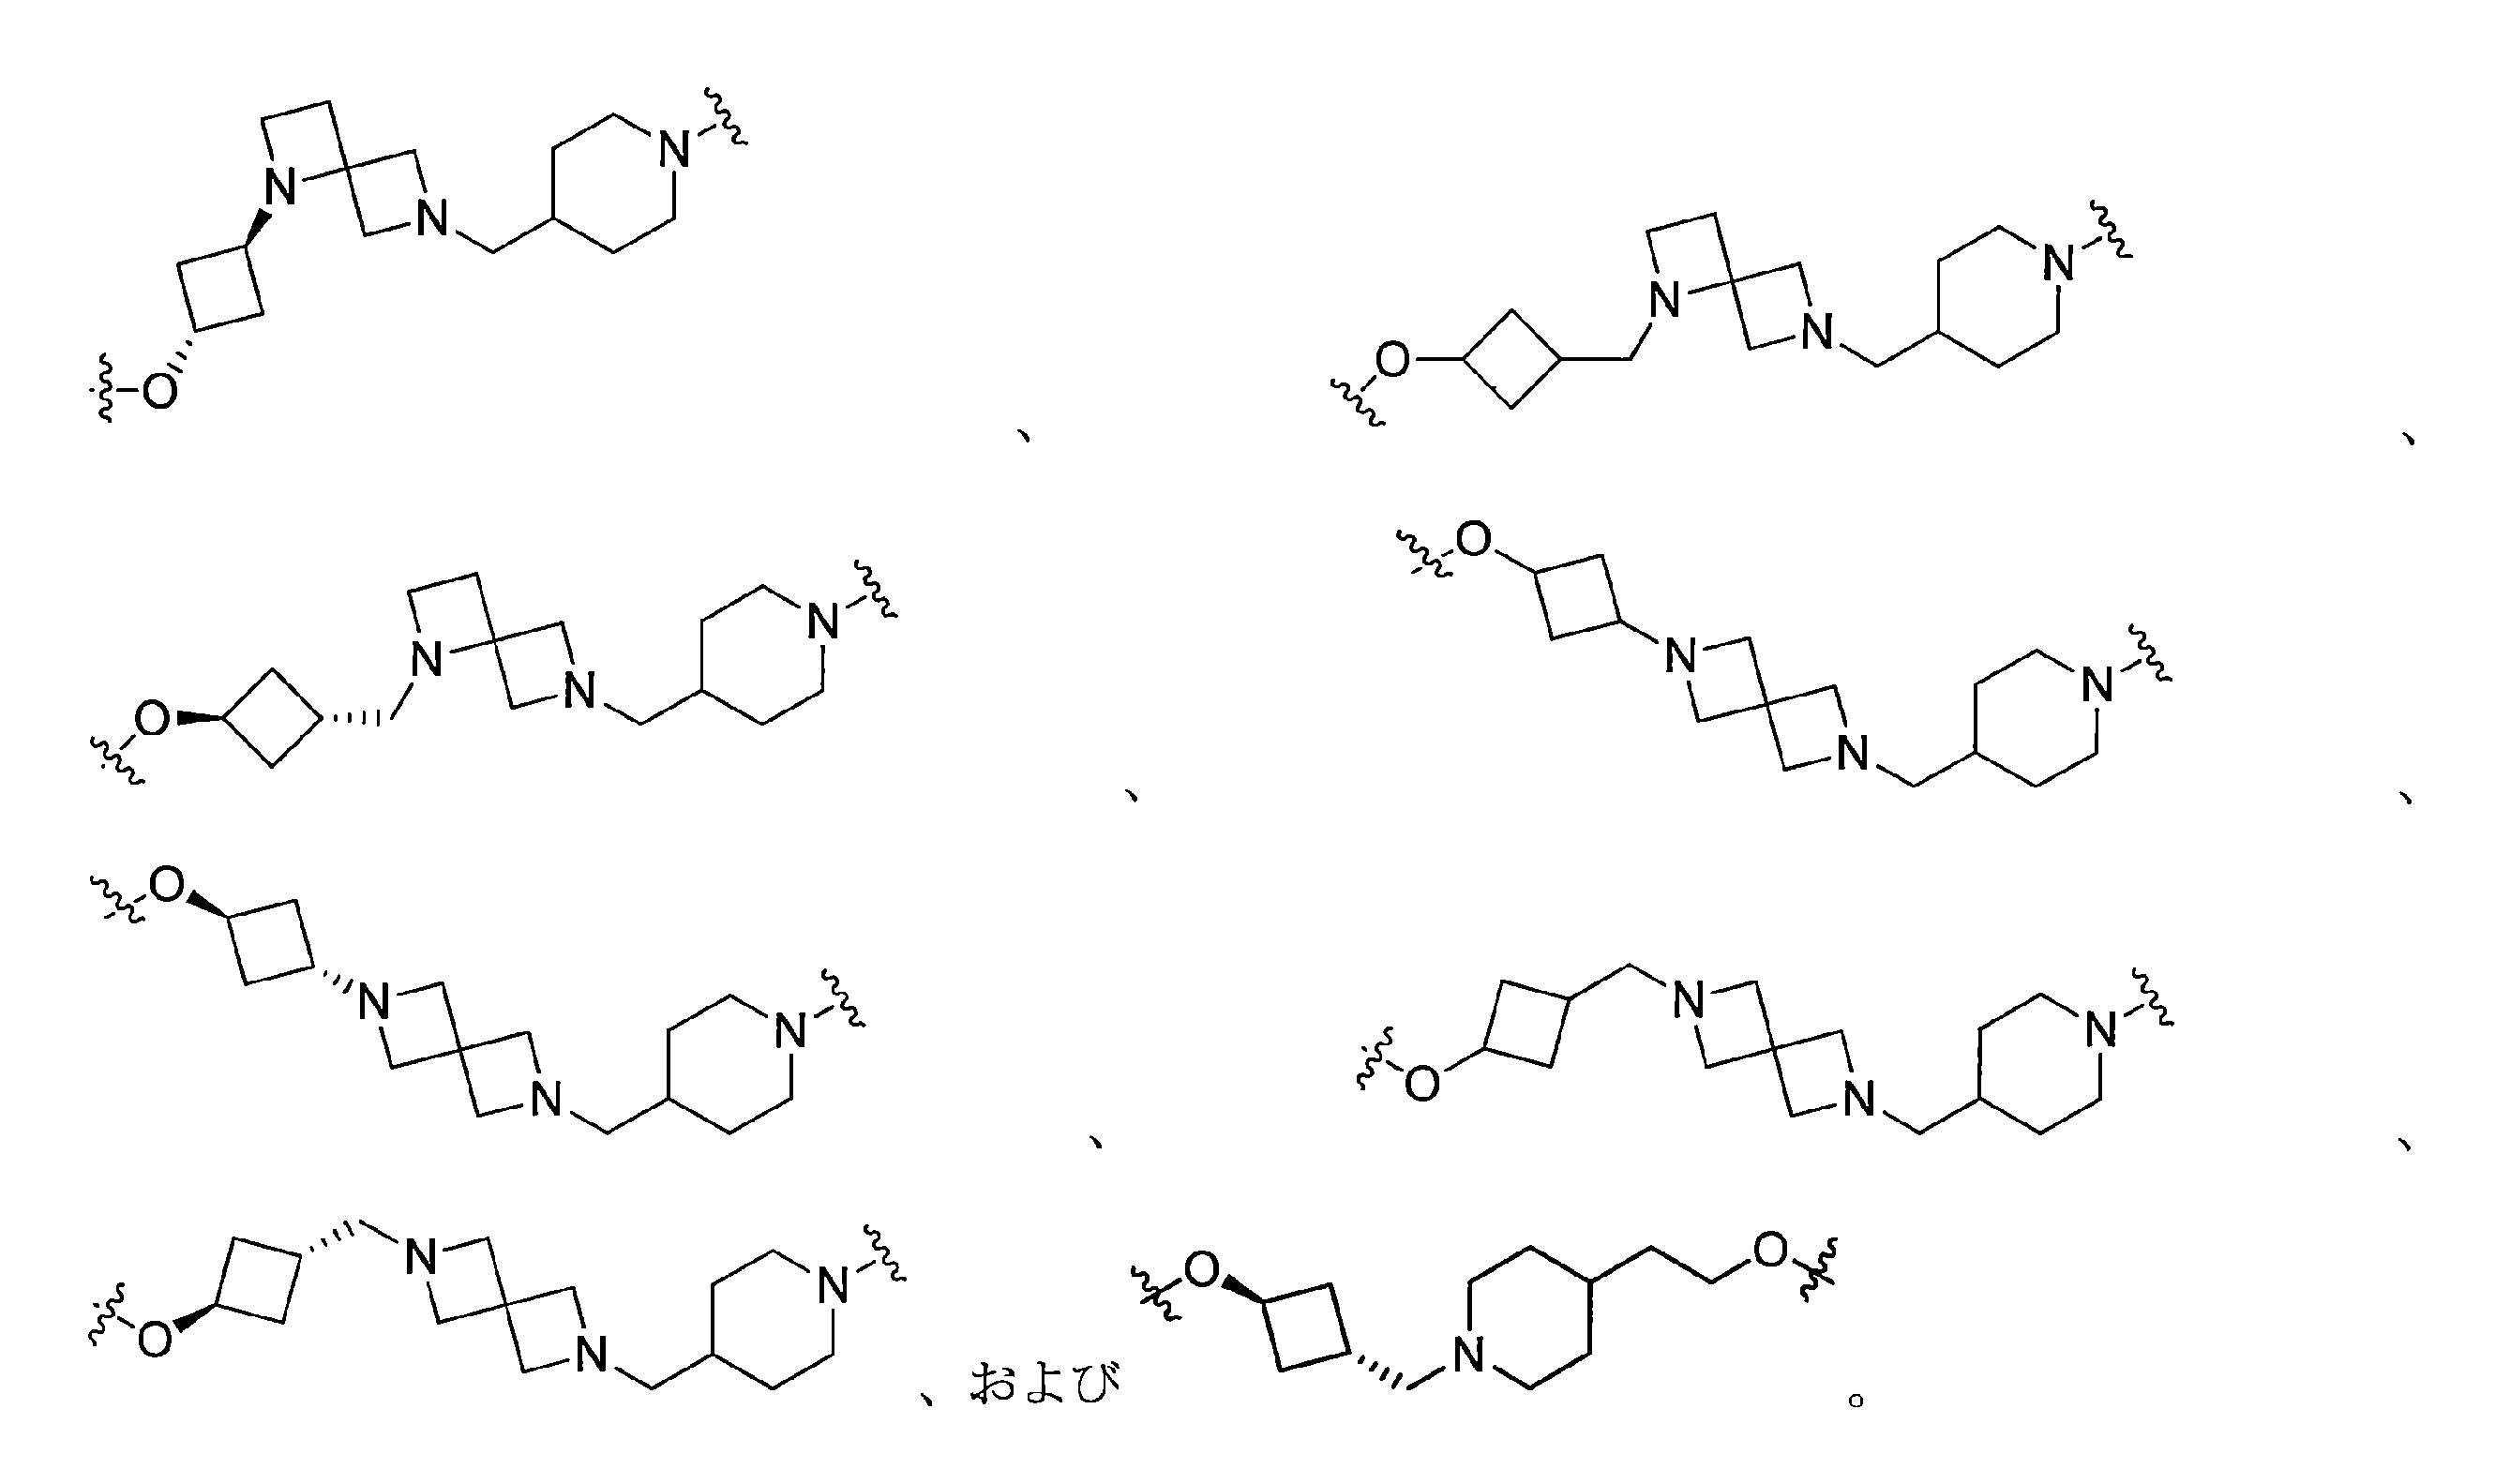
本明細書に記載される任意の態様または実施形態において、リンカー(L)は、以下からなる群から選択される:

追加的実施形態において、リンカー(L)は、限定されないが以下に示される構造から選択される構造を含み、ここで破線はPTM部分またはULM部分への結合点を示す:
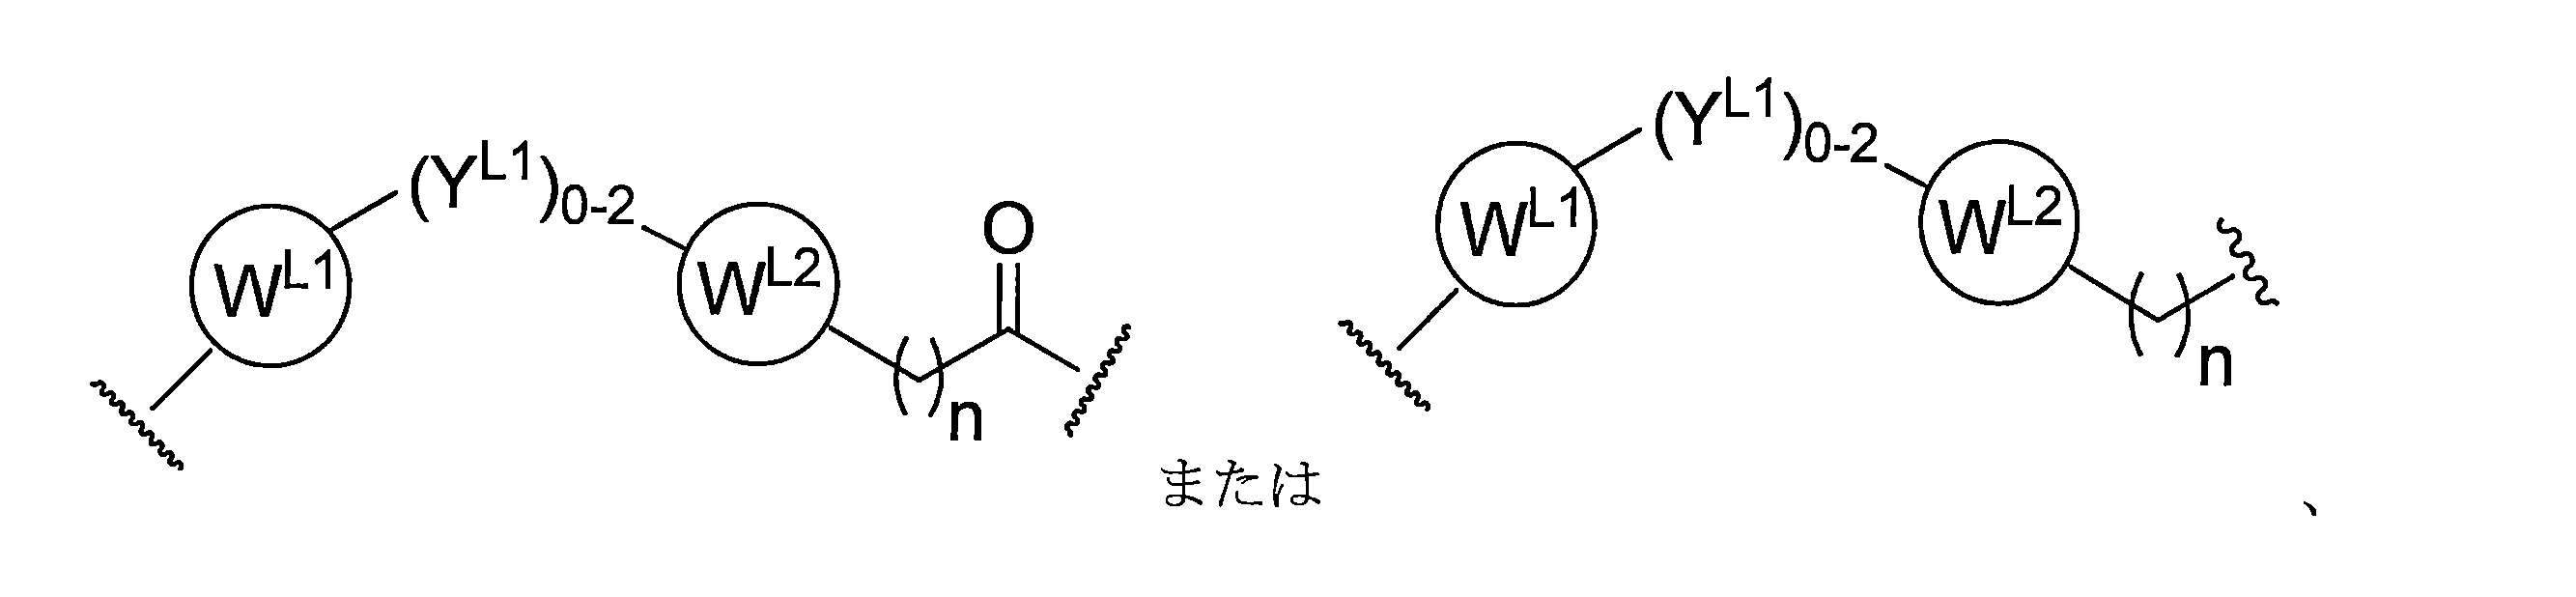
WL1およびWL2は、各々独立して、存在しないか、任意にRQで置換された0~4個のヘテロ原子を有する4~8員であり、各RQは、独立して、H、ハロ、OH、CN、CF3、任意に置換された直鎖もしくは分枝鎖C1-C6アルキル、任意に置換された直鎖もしくは分枝鎖C1-C6アルコキシであるか、または2個のRQ基は、それらが結合している原子と一緒になって、0~4個のヘテロ原子を含有する4~8員環系を形成し、
YL1は、各々独立して、結合、(直鎖、分岐鎖、任意に置換された)かつ任意に1個以上のC原子がOで置き換えられたC1-C6アルキル(されている)、または(直鎖、分岐鎖、任意に置換された)C1-C6アルコキシであり、
nは、0~10であり、

WL1 and WL2 are each independently 4-8 membered, absent or having 0-4 heteroatoms optionally substituted with RQ, each RQ is independently H, halo, OH, CN, CF3, optionally substituted straight or branched chain C1-C6 alkyl, optionally substituted straight or branched chain C1-C6 alkoxy, or two RQ groups taken together with the atom to which they are attached form a 4-8 membered ring system containing 0-4 heteroatoms;
Each YL1 is independently a bond, a C1-C6 alkyl (linear, branched, optionally substituted) and optionally one or more C atoms are replaced by O, or a C1-C6 alkoxy (linear, branched, optionally substituted);
n is 0 to 10;
追加的実施形態において、リンカー(L)は、限定されないが以下に示される構造から選択される構造を含み、ここで破線はPTM部分またはULM部分への結合点を示す:
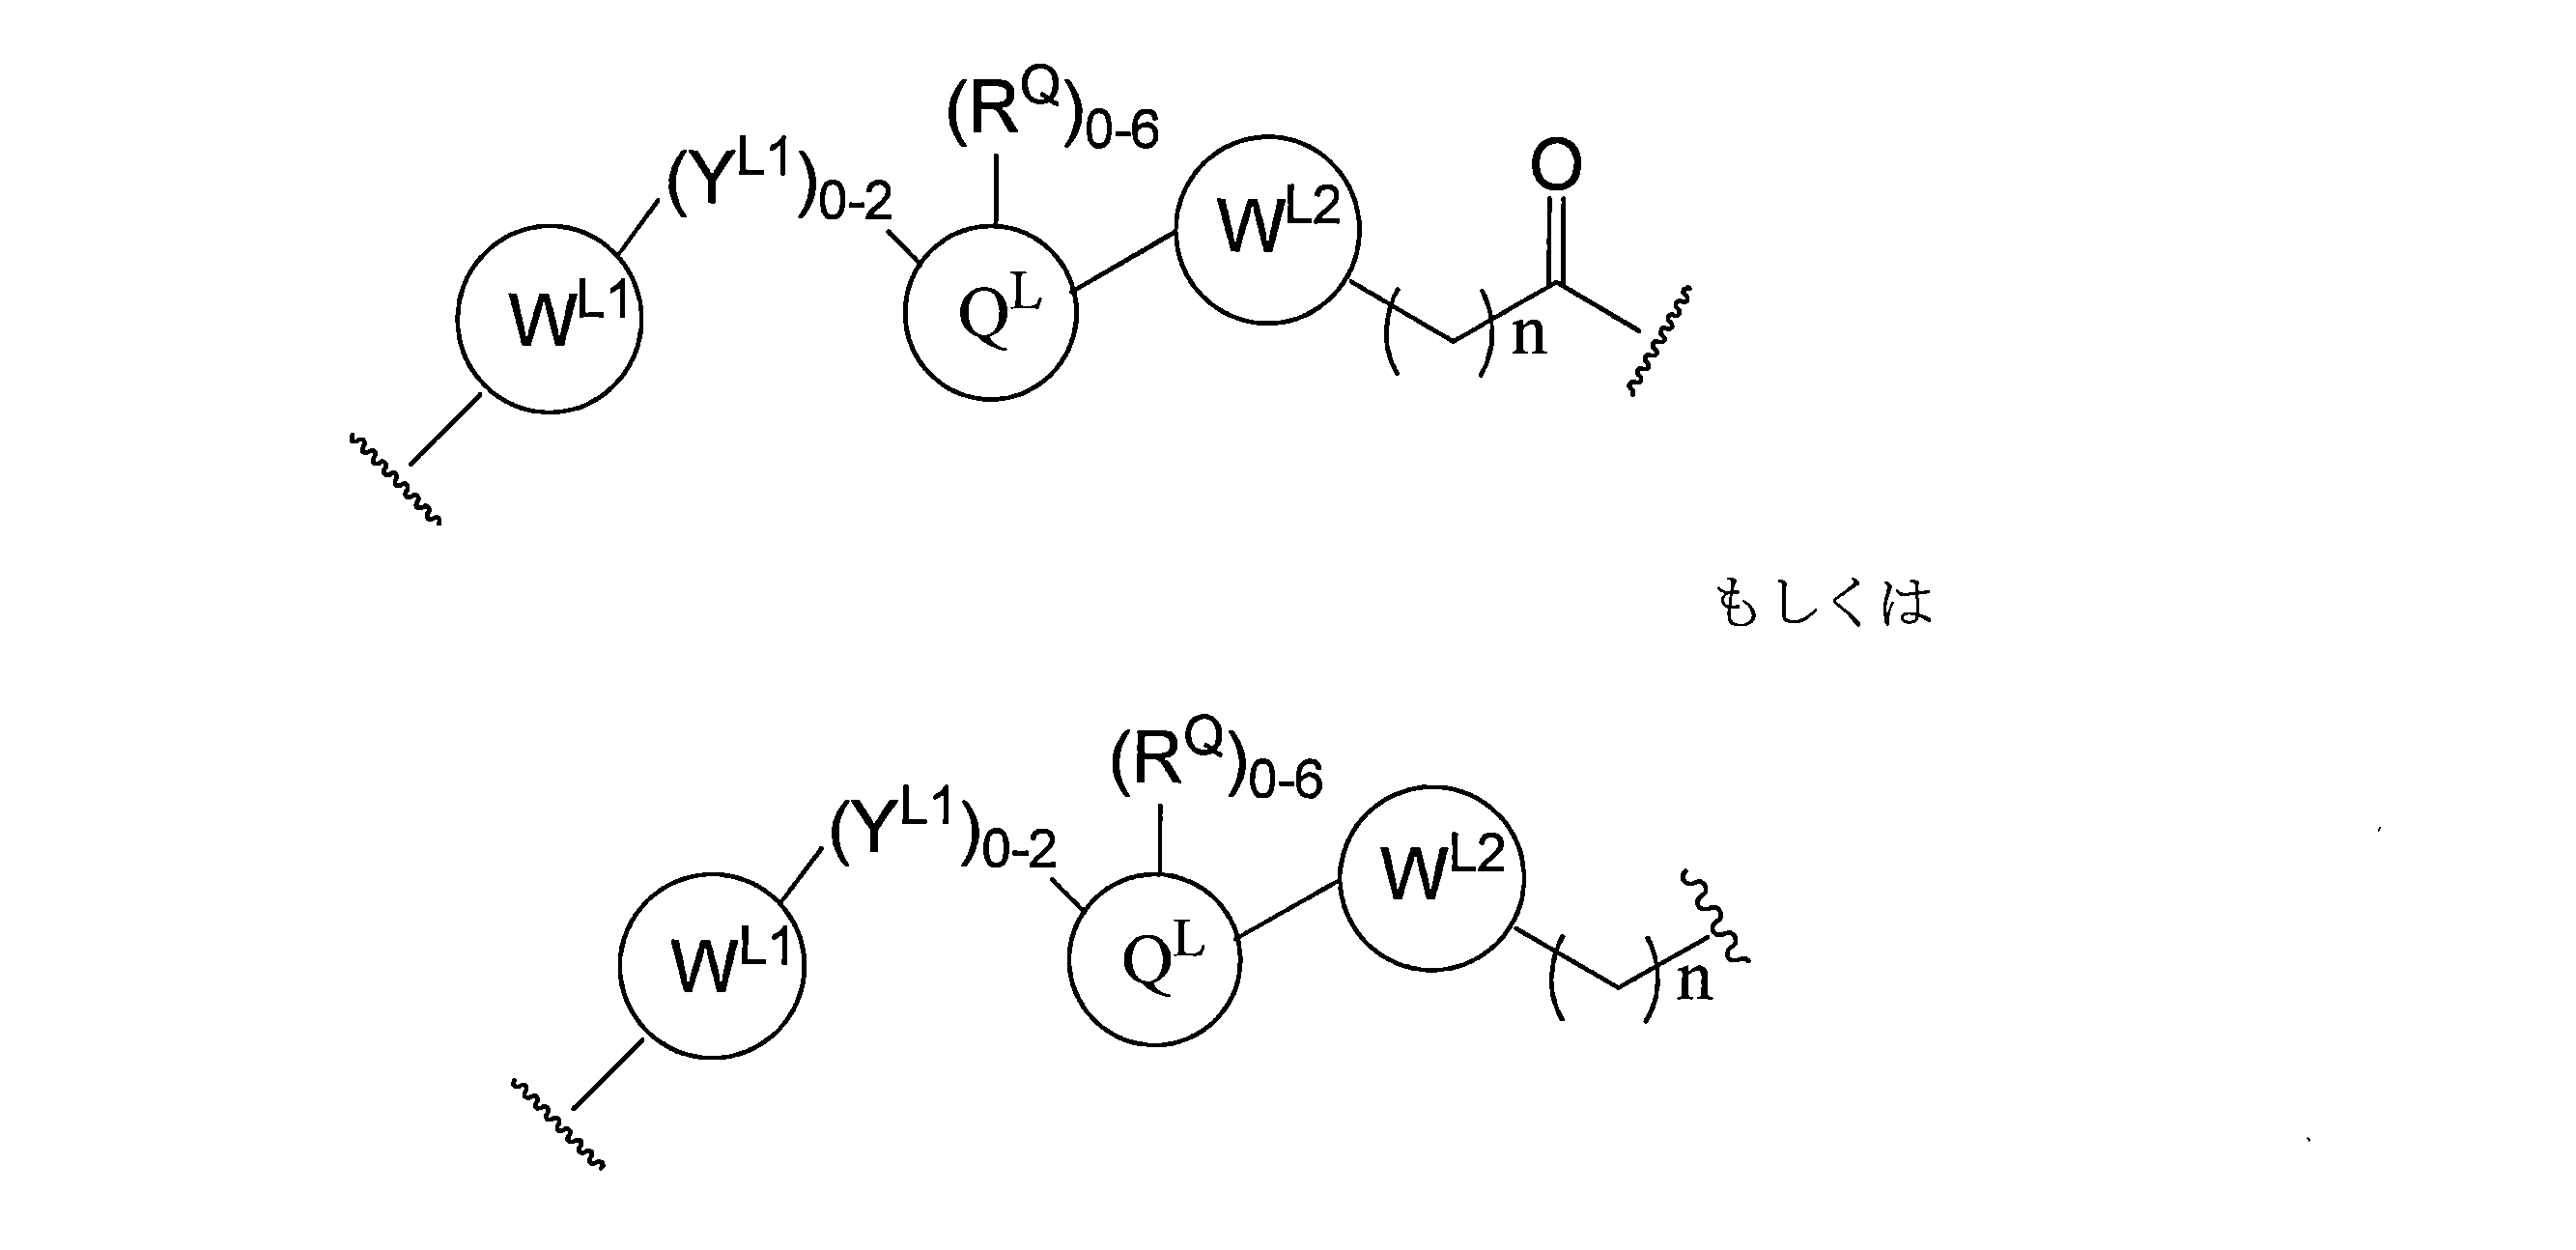
WL1およびWL2は、各々独立して、存在しないか、アリール、ヘテロアリール、環式、ヘテロシクリル、任意に1個以上のC原子がOもしくはNで置き換えられたC1-6アルキル、任意に1個以上のC原子がOで置き換えられたC1-6アルケニル、任意に1個以上のC原子がOで置き換えられたC1-6アルキニル、二環式、ビアリール、ビヘテロアリール、またはビヘテロシクリルであり、各々が、RQで任意に置換され、各RQは、独立して、H、ハロ、OH、CN、CF3、ヒドロキシル、ニトロ、C≡CH、C2-6アルケニル、C2-6アルキニル、任意に置換された直鎖または分岐鎖C1-C6アルキル、任意に置換された直鎖または分岐鎖C1-C6アルコキシ、任意に置換されたOC1-3アルキル(例えば、1個以上のFで任意に置換されている)、OH、NH2、NRY1RY2、CNであるか、または2つのRQ基が、それらが結合している原子と一緒になって、0~4個のヘテロ原子を含有する4~8員環系を形成し、
YL1は、各々独立して、結合、NRYL1、O、S、NRYL2、CRYL1RYL2、C=O、C=S、SO、SO2、(直鎖、分岐鎖、任意に置換された)かつ任意に1個以上のC原子がOで置き換えられたC1-C6アルキル、または(直鎖、分岐鎖、任意に置換された)C1-C6アルコキシであり、
QLは、任意に架橋され、任意に0~6個のRQで置換された、0~4個のヘテロ原子を有する3~6員の脂環式または芳香環であり、各RQは、独立して、H、任意に1つ以上のハロもしくはC1-6アルコキシルで置換された、直鎖もしくは分岐鎖C1-6アルキルであるか、または2個のRQ基は、それらが結合している原子と一緒になって、0~2個のヘテロ原子を含有する3~8員環系を形成し、
RYL1、RYL2は、各々独立して、H、OH、(直鎖、分岐鎖、任意に1つ以上のハロ、C1-6アルコキシルで置換された)C1-6アルキルであるか、またはR1、R2は、それらが結合している原子と一緒になって、0~2個のヘテロ原子を含有する3~8員環系を形成し、
nは、0~10であり、
WL1 and WL2 are each independently absent, aryl, heteroaryl, cyclic, heterocyclyl, C1-6 alkyl optionally having one or more C atoms replaced by O or N, C1-6 alkenyl optionally having one or more C atoms replaced by O, C1-6 alkynyl optionally having one or more C atoms replaced by O, bicyclic, biaryl, biheteroaryl, or biheterocyclyl, each optionally substituted with RQ, and each RQ is independently H, halo, OH, C N, CF3, hydroxyl, nitro, C≡CH, C2-6 alkenyl, C2-6 alkynyl, optionally substituted straight or branched chain C1-C6 alkyl, optionally substituted straight or branched chain C1-C6 alkoxy, optionally substituted OC1-3 alkyl (e.g. optionally substituted with one or more F), OH, NH2, NRY1RY2, CN, or two RQ groups together with the atom to which they are attached form a 4-8 membered ring system containing 0-4 heteroatoms;
Each YL1 is independently a bond, NRYL1, O, S, NRYL2, CRYL1RYL2, C=O, C=S, SO, SO2, C1-C6 alkyl (linear, branched, optionally substituted) and optionally one or more C atoms are replaced by O, or C1-C6 alkoxy (linear, branched, optionally substituted);
QL is an optionally bridged 3-6 membered alicyclic or aromatic ring having 0-4 heteroatoms, optionally substituted with 0-6 RQ, each RQ is independently H, a straight or branched C1-6 alkyl optionally substituted with one or more halo or C1-6 alkoxyl, or two RQ groups together with the atom to which they are attached form a 3-8 membered ring system containing 0-2 heteroatoms;
RYL1, RYL2 are each independently H, OH, C1-6 alkyl (linear, branched, optionally substituted with one or more halo, C1-6 alkoxyl), or R1, R2 together with the atom to which they are attached form a 3-8 membered ring system containing 0-2 heteroatoms;
n is 0 to 10;
追加的実施形態では、リンカー基は、約1~約100個のエチレングリコール単位、約1~約50個のエチレングリコール単位、約1~約25個のエチレングリコール単位、約1~10個のエチレングリコール単位、1~約8個のエチレングリコール単位、および1~6個のエチレングリコール単位、約2~4個のエチレングリコール単位を有する任意に置換された(ポリ)エチレングリコールであるか、または任意に置換されたO、N、S、PまたはSi原子が散在する任意に置換されたアルキル基である。ある特定の実施形態では、リンカーは、アリール、フェニル、ベンジル、アルキル、アルキレン、またはヘテロシクリル基で置換されている。特定の実施形態では、リンカーは非対称または対称であってもよい。 In additional embodiments, the linker group is an optionally substituted (poly)ethylene glycol having about 1 to about 100 ethylene glycol units, about 1 to about 50 ethylene glycol units, about 1 to about 25 ethylene glycol units, about 1 to 10 ethylene glycol units, 1 to about 8 ethylene glycol units, and 1 to 6 ethylene glycol units, about 2 to 4 ethylene glycol units, or an optionally substituted alkyl group interspersed with optionally substituted O, N, S, P, or Si atoms. In certain embodiments, the linker is substituted with an aryl, phenyl, benzyl, alkyl, alkylene, or heterocyclyl group. In certain embodiments, the linker may be asymmetric or symmetric.
本明細書に記載の化合物の実施形態のいずれかにおいて、リンカー基は、本明細書に記載される任意の適切な部分であってもよい。1つの実施形態では、リンカーは、約1~約12エチレングリコール単位、1~約10エチレングリコール単位、約2~6エチレングリコール単位、約2~5エチレングリコール単位、約2~4エチレングリコール単位のサイズ範囲の置換または非置換のポリエチレングリコール基である。 In any of the embodiments of the compounds described herein, the linker group may be any suitable moiety described herein. In one embodiment, the linker is a substituted or unsubstituted polyethylene glycol group ranging in size from about 1 to about 12 ethylene glycol units, from 1 to about 10 ethylene glycol units, from about 2 to 6 ethylene glycol units, from about 2 to 5 ethylene glycol units, from about 2 to 4 ethylene glycol units.
別の実施形態では、本開示は、上記のようなPTM基を含む化合物を対象とし、PTM基は、ユビキチンリガーゼによってユビキチン化され、かつULM基に直接またはリンカー部分Lを介して化学結合された、標的タンパク質またはポリペプチド(例えば、SMARCA2、BRAHMA、またはBRM)に結合するか、あるいはPTMが、代替的に、これもまたユビキチンリガーゼ結合部分であるULM’基であり、ULM’基は、上記のようなULM基と同じであっても異なっていてもよく、Lは、上記のようなリンカー部分であり、リンカー部分は存在しても存在しなくてもよく、かつULMをPTM、またはその薬学的に許容される塩、鏡像異性体、立体異性体、溶媒和物、もしくは多形に化学(共有)結合する。 In another embodiment, the disclosure is directed to compounds comprising a PTM group as described above, which is ubiquitinated by a ubiquitin ligase and is attached to a target protein or polypeptide (e.g., SMARCA2, BRAHMA, or BRM) that is chemically attached to a ULM group, either directly or via a linker moiety L, or the PTM is alternatively a ULM' group that is also a ubiquitin ligase binding moiety, which may be the same or different as the ULM group as described above, L is a linker moiety as described above, which may or may not be present, and which chemically (covalently) attaches the ULM to the PTM, or a pharma- ceutically acceptable salt, enantiomer, stereoisomer, solvate, or polymorph thereof.
特定の実施形態では、リンカー基Lは、以下からなる群から独立して選択される1つ以上の共有結合された構造単位を含む基である:

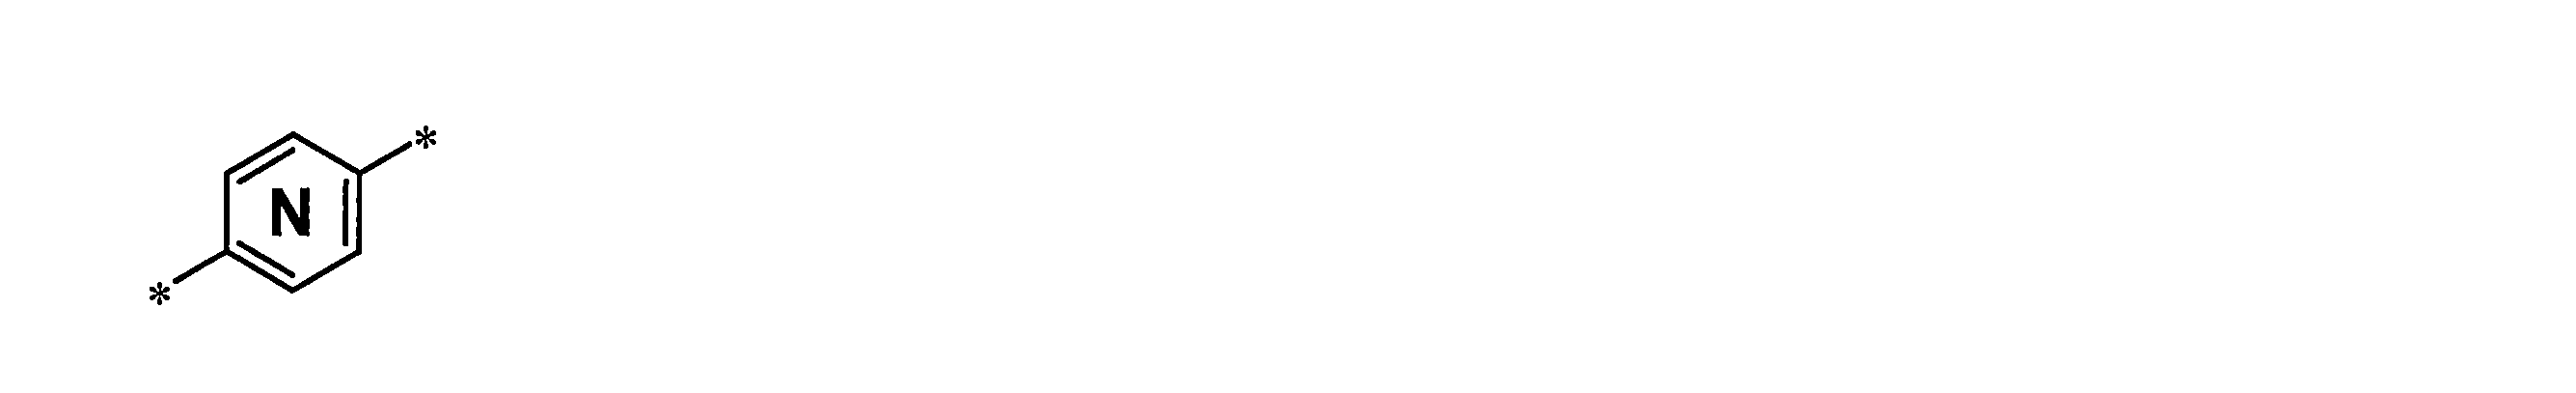
ULM基とPTM基は、リンカーの化学的性質に対し適切であり、安定的な任意の基を介してリンカー基に共有結合されてもよく、本開示の好ましい態様においては、リンカーは独立してULM基とPTM基に、好ましくはアミド、エステル、チオエステル、ケト基、カルバミン酸塩(ウレタン)、炭素またはエーテルを介して共有結合され、それら各基は、ULM基とPTM基上のいずれかに挿入され、ユビキチンリガーゼに対するULM基の最大結合と、分解される標的タンパク質に対するPTM基の最大結合を提供し得る。(PTM基がULM基である特定の態様では、分解される標的タンパク質は、ユビキチンリガーゼそれ自体でもあり得ることに留意されたい)。ある特定の態様では、リンカーは、ULMおよび/またはPTM基上の、任意に置換されたアルキル、アルキレン、アルケニル、もしくはアルキニル基、アリール基、またはヘテロシクリル基に結合され得る。 The ULM and PTM groups may be covalently attached to the linker group via any group that is appropriate and stable for the linker chemistry, and in preferred embodiments of the present disclosure, the linker is independently covalently attached to the ULM and PTM groups, preferably via an amide, ester, thioester, keto group, carbamate (urethane), carbon, or ether, each of which may be inserted either on the ULM or PTM group to provide maximum binding of the ULM group to the ubiquitin ligase and maximum binding of the PTM group to the target protein to be degraded. (Note that in certain embodiments where the PTM group is a ULM group, the target protein to be degraded may also be the ubiquitin ligase itself). In certain embodiments, the linker may be attached to an optionally substituted alkyl, alkylene, alkenyl, or alkynyl group, aryl group, or heterocyclyl group on the ULM and/or PTM group.
例示的なPTM
本開示の任意の態様または実施形態では、PTM基は、標的タンパク質、例えば、スイッチ/スクロース非発酵性(SWI/SNF)関連、マトリックス関連、アクチン依存性調節因子のクロマチン、サブファミリーA、メンバー2(SMARCA2)またはBRMに結合する、部分である。したがって、本明細書に記載される任意の態様または実施形態では、PTM基は、SMARCA2またはBRMタンパク質に特異的に結合する(標的タンパク質SMARCA2、BRAHMAまたはBRMに結合する)任意の部分である。
Exemplary PTMs
In any aspect or embodiment of the present disclosure, the PTM group is a moiety that binds to a target protein, e.g., switch/sucrose non-fermenting (SWI/SNF)-associated, matrix-associated, actin-dependent regulator of chromatin, subfamily A, member 2 (SMARCA2) or BRM. Thus, in any aspect or embodiment described herein, the PTM group is any moiety that specifically binds to a SMARCA2 or BRM protein (binds to the target protein SMARCA2, BRAHMA or BRM).
ある特定の実施形態では、本明細書に記載される化合物は、標的タンパク質、例えば、Brmに結合する手段を含む。したがって、ある特定の態様では、本開示は、Brmに結合するための手段、およびVHLに結合するための手段、ならびにBrmに結合するための手段をVHLに結合するための手段に化学的に結合するための手段を有する二官能性化合物を提供する。 In certain embodiments, the compounds described herein include a means for binding to a target protein, e.g., Brm. Thus, in certain aspects, the present disclosure provides bifunctional compounds having a means for binding to Brm and a means for binding to VHL, as well as a means for chemically linking the means for binding to Brm to the means for binding to VHL.
以下に記載される組成物は、一連の低分子標的タンパク質結合部分のいくつかを例示する。また、このような小分子標的タンパク質結合部分には、これらの組成物の薬学的に許容される塩、鏡像異性体、溶媒和物、および多形、ならびにSMARCA2を標的化し得る他の小分子も含まれる。これらの結合部分は、ユビキチンリガーゼに近接してユビキチン化および分解を行う目的で、標的タンパク質(そこにタンパク質標的部分が結合される)を提示するために、好ましくはリンカーを通してユビキチンリガーゼ結合部分に結合される。タンパク質標的部分またはPTM基に結合し、かつユビキチンリガーゼに作用するか、またはユビキチンリガーゼによって分解することができる、任意のタンパク質(例えば、SMARCA2、BRAHMA、またはBRM)は、本開示に従う標的タンパク質である。 The compositions described below exemplify some of the series of small molecule target protein binding moieties. Such small molecule target protein binding moieties also include pharma- ceutically acceptable salts, enantiomers, solvates, and polymorphs of these compositions, as well as other small molecules that may target SMARCA2. These binding moieties are attached to the ubiquitin ligase binding moiety, preferably through a linker, to present the target protein (to which the protein target moiety is attached) in proximity to the ubiquitin ligase for ubiquitination and degradation. Any protein (e.g., SMARCA2, BRAHMA, or BRM) that is attached to a protein target moiety or PTM group and can act on or be degraded by a ubiquitin ligase is a target protein in accordance with the present disclosure.
本開示を使用して、タンパク質が調節不全(例えば、SMARCA4欠損/変異)であり、かつ患者がSMARCA2、BRAHMA、またはBRMなどのタンパク質の分解および/または阻害から利益を得るであろう任意の疾患状態および/または状態を含む、多数の疾患状態および/または病態を治療し得る。 The present disclosure may be used to treat numerous disease states and/or conditions, including any disease state and/or condition in which a protein is dysregulated (e.g., SMARCA4 deficiency/mutation) and in which the patient would benefit from degradation and/or inhibition of a protein such as SMARCA2, BRAHMA, or BRM.
追加の態様において本明細書は、本明細書に記載の化合物またはその塩型の有効量、および薬学的に許容可能な担体、添加剤または賦形剤、および任意で追加の生物活性剤を含む治療用組成物を提供する。治療用組成物は、患者または対象、例えばヒトなどの動物においてタンパク質の分解を調節し、当該分解されたタンパク質を介して調節される病状もしくは状態の治療または改善に使用することができる。ある特定の実施形態では、本明細書に記載される治療組成物を使用して、肺がんまたは非小細胞肺がんを含む、疾患、例えば、SWI/SNF関連がん、SMARCA4変異関連がん、SMARCA4欠損がん、または通常のSMARCA4発現と比べてSMARCA4の発現が減少した(例えば、野生型SMARCA4を有する同様の位置にある非がん性細胞における非変異SMARCA4またはSMARCA4の発現と比べて発現が減少した)がんのうちの少なくとも1つなどのがんの治療または改善のために、目的のタンパク質の分解をもたらし得る。本明細書に記載される任意の態様または実施形態では、がんは、SWI/SNF関連がん、SMARCA4変異を有するがん、SMARCA4欠損を有するがん、またはそれらの組み合わせのうちの少なくとも1つであり、肺がんまたは非小細胞肺がんであり得る。 In a further aspect, the present disclosure provides a therapeutic composition comprising an effective amount of a compound or salt form thereof as described herein, and a pharma- ceutically acceptable carrier, additive or excipient, and optionally an additional bioactive agent. The therapeutic composition can be used to modulate protein degradation in a patient or subject, such as an animal, such as a human, and to treat or ameliorate a disease state or condition regulated via the degraded protein. In certain embodiments, the therapeutic composition described herein can be used to effect degradation of a protein of interest for the treatment or amelioration of a disease, such as at least one of SWI/SNF-associated cancer, SMARCA4 mutation-associated cancer, SMARCA4-deficient cancer, or cancer with reduced expression of SMARCA4 compared to normal SMARCA4 expression (e.g., reduced expression compared to non-mutated SMARCA4 or expression of SMARCA4 in a similarly located non-cancerous cell with wild-type SMARCA4), including lung cancer or non-small cell lung cancer. In any aspect or embodiment described herein, the cancer is at least one of a SWI/SNF-associated cancer, a SMARCA4-mutated cancer, a SMARCA4-deficient cancer, or a combination thereof, and may be lung cancer or non-small cell lung cancer.
ある特定の追加の実施形態では、本明細書に記載される治療組成物を使用して、疾患、例えば、SWI/SNF関連がん、SMARCA2関連がん、またはSMARCA2の正常発現もしくは過剰発現を有するがんのうちの少なくとも1つなどのがんの治療または改善のために、目的のタンパク質の分解をもたらし得る。 In certain additional embodiments, the therapeutic compositions described herein may be used to effect degradation of a protein of interest for the treatment or amelioration of a disease, such as at least one of a SWI/SNF-associated cancer, a SMARCA2-associated cancer, or a cancer having normal or overexpression of SMARCA2.
別の態様では、本開示は、タンパク質またはポリペプチドを分解することにより、その必要のある対象の病態を治療する方法、または疾患もしくは状態の症状を改善する方法に関するものであり、ここで当該タンパク質またはポリペプチドを介して病態または状態は調節されており、当該方法は、該患者または対象に、上述の少なくとも1つの化合物の有効量、例えば治療有効量を、任意で薬学的に許容可能な担体、添加剤または賦形剤、および任意で追加の生物活性剤と併用して投与することを含み、ここで当該組成物は、当該対象の疾患もしくは障害もしくはその症状の治療または改善に有効である。本開示方法を使用して、本明細書に記載される少なくとも1つの化合物の有効量を投与することにより、がんを含む多くの症状または状態が治療され得る。病態または状態は、微生物体、または例えばウイルス、細菌、真菌、原虫もしくはその他の微生物などの他の外来性主体により引き起こされる疾患であってもよく、または病態および/もしくは状態が生じるタンパク質の過剰発現により引き起こされる病態であってもよい。 In another aspect, the disclosure relates to a method of treating a pathology or ameliorating a symptom of a disease or condition in a subject in need thereof by degrading a protein or polypeptide, where the pathology or condition is regulated via the protein or polypeptide, the method comprising administering to the patient or subject an effective amount, e.g., a therapeutically effective amount, of at least one compound as described above, optionally in combination with a pharma- ceutically acceptable carrier, additive or excipient, and optionally an additional bioactive agent, where the composition is effective to treat or ameliorate the disease or disorder or a symptom thereof in the subject. Using the disclosed method, many conditions or symptoms can be treated, including cancer, by administering an effective amount of at least one compound described herein. The pathology or condition may be a disease caused by a microbial organism or other foreign agent, such as, for example, a virus, bacteria, fungus, protozoa, or other microorganism, or may be a pathology caused by overexpression of a protein resulting in the pathology and/or condition.
別の態様において本明細書は、本開示による化合物を使用して生物的システムにおいて対象タンパク質の分解効果を特定する方法を提供する。 In another aspect, the present specification provides a method for identifying the degradative effect of a protein of interest in a biological system using a compound according to the present disclosure.
「標的タンパク質」という用語は、以下の本明細書において、本開示化合物が結合し、ユビキチンリガーゼにより分解される標的であるタンパク質またはポリペプチドを記述するために使用される。かかる低分子標的タンパク質結合部分には、これらの組成物の薬学的に許容可能な塩、鏡像異性体、溶媒和物、および多形体、ならびに対象のタンパク質を標的としうる他の低分子も含まれる。これらの結合部分は、少なくとも1つのリンカー基Lを介して、少なくとも1つのULM基(例えば、VLM)に結合される。 The term "target protein" is used herein below to describe a protein or polypeptide that is a target to which the disclosed compounds bind and are degraded by ubiquitin ligase. Such small molecule target protein binding moieties also include pharma- ceutically acceptable salts, enantiomers, solvates, and polymorphs of these compositions, as well as other small molecules that may target a protein of interest. These binding moieties are attached to at least one ULM group (e.g., VLM) via at least one linker group L.
このタンパク質標的は、当該タンパク質に結合する化合物部分を特定するスクリーニングにおいて使用されてもよく、そして本開示化合物内に当該部分を組み込むことによって、治療の最終結果に対する、当該タンパク質の活性レベルを変化させることができる。 This protein target may be used in screens to identify compound moieties that bind to the protein, and by incorporating the moieties into the disclosed compounds, the activity level of the protein can be altered for therapeutic end results.
「タンパク質標的部分」またはPTMという用語は、標的タンパク質、またはSMARCA2もしくはBRMなどの他の目的のタンパク質もしくはポリペプチドに結合し、ユビキチンリガーゼによるタンパク質またはポリペプチドの分解が生じ得るように、そのタンパク質またはポリペプチドをユビキチンリガーゼに近接して配置/提示する、小分子を説明するために使用される。以下に記載される組成物は、低分子標的タンパク質の構成因子のうちのいくつかを例示する。 The term "protein targeting moiety" or PTM is used to describe a small molecule that binds to a target protein or other protein or polypeptide of interest, such as SMARCA2 or BRM, and positions/presents the protein or polypeptide in proximity to a ubiquitin ligase so that degradation of the protein or polypeptide by the ubiquitin ligase can occur. The compositions described below exemplify some of the components of small molecule target proteins.
本明細書に記載される任意の態様または実施形態では、本開示のPTMは、以下によって表される化学構造を有し、

WPTM1は、任意に置換された5~6員アリールまたはヘテロアリール環(例えば、ヒドロキシ、ハロゲン、アルコキシ、アルキル、ハロアルキル、ホスフェート、アミノ、アルキルアミノ、シアノ、またはそれらの組み合わせから選択される0、1、2、または3個の置換基で置換された5~6員アリールまたはヘテロアリール)であり、
WPTM2は、任意に置換された5~6員アリールまたはヘテロアリール環(例えば、ヒドロキシ、ハロゲン、アルコキシ、アルキル、ハロアルキル、アミノ、アルキルアミノ、およびシアノから選択される0、1、2、または3個の置換基で置換された5~6員アリールまたはヘテロアリール)であり、
WPTM3は、任意に置換された3~9員アリールもしくはヘテロアリール環(例えば、任意に置換された5~6員アリールもしくはヘテロアリール環、またはヒドロキシ、ハロゲン、アルコキシ、アルキル、ハロアルキル、アミノ、アルキルアミノ、およびシアノから選択される0、1、2、もしくは3個の置換基で置換された3~9もしくは5~6員アリールもしくはヘテロアリール)、または任意に置換された4~9員シクロアルキルもしくはヘテロシクリル、例えば、任意に置換された架橋ビシクロアルキルおよび架橋ビヘテロシクリル環(例えば、ヒドロキシ、ハロゲン、アルコキシ、アルキル、ハロアルキル、アミノ、アルキルアミノ、およびシアノから選択される0、1、もしくは2個の置換基で置換された4~9員シクロアルキルもしくはヘテロシクリル)であり、
WPTM5は、存在しない(例えば、WPTM3が、L(リンカー)またはULMに直接連結されている)か、または任意に置換されたアルキル、任意に置換された5~6員シクロアルキル、複素環、アリール、もしくはヘテロアリール環(例えば、ヒドロキシ、ハロゲン、アルコキシ、アルキル、ハロアルキル、アミノ、アルキルアミノ、およびシアノから選択される0、1、または2個の置換基で置換された、5~6員シクロアルキル、複素環、アリール、またはヘテロアリール)であり、

WPTM1 is an optionally substituted 5-6 membered aryl or heteroaryl ring (e.g., a 5-6 membered aryl or heteroaryl substituted with 0, 1, 2, or 3 substituents selected from hydroxy, halogen, alkoxy, alkyl, haloalkyl, phosphate, amino, alkylamino, cyano, or combinations thereof);
WPTM2 is an optionally substituted 5-6 membered aryl or heteroaryl ring (e.g., a 5-6 membered aryl or heteroaryl substituted with 0, 1, 2, or 3 substituents selected from hydroxy, halogen, alkoxy, alkyl, haloalkyl, amino, alkylamino, and cyano);
WPTM3 is an optionally substituted 3-9 membered aryl or heteroaryl ring (e.g., an optionally substituted 5-6 membered aryl or heteroaryl ring, or a 3-9 or 5-6 membered aryl or heteroaryl substituted with 0, 1, 2, or 3 substituents selected from hydroxy, halogen, alkoxy, alkyl, haloalkyl, amino, alkylamino, and cyano), or an optionally substituted 4-9 membered cycloalkyl or heterocyclyl, such as optionally substituted bridged bicycloalkyl and bridged biheterocyclyl rings (e.g., a 4-9 membered cycloalkyl or heterocyclyl substituted with 0, 1, or 2 substituents selected from hydroxy, halogen, alkoxy, alkyl, haloalkyl, amino, alkylamino, and cyano);
WPTM5 is absent (e.g., WPTM3 is directly linked to L (linker) or ULM) or is an optionally substituted alkyl, an optionally substituted 5-6 membered cycloalkyl, heterocycle, aryl, or heteroaryl ring (e.g., a 5-6 membered cycloalkyl, heterocycle, aryl, or heteroaryl substituted with 0, 1, or 2 substituents selected from hydroxy, halogen, alkoxy, alkyl, haloalkyl, amino, alkylamino, and cyano);
本明細書に記載される任意の態様または実施形態では、WPTM5は、ピペリジンである。 In any aspect or embodiment described herein, WPTM5 is piperidine.
ある特定の実施形態では、WPTM1は、ホスフェート置換を含む。 In certain embodiments, WPTM1 includes a phosphate substitution.
本明細書に記載される任意の態様または実施形態では、本開示のPROTACのPTMは、式Iによって表され、式中、
WPTM1は、任意に置換されたフェニルまたはピリジル(例えば、本明細書に記載されるように置換された、例えば、本明細書に記載されるように選択される追加の任意の置換基を有するかまたは有しないヒドロキシまたはホスフェート置換基で置換された、例えば、ヒドロキシ、ハロゲン、アルコキシ、アルキル、ハロアルキル、アミノ、アルキルアミノ、シアノ、またはそれらの組み合わせから選択される0、1、2、または3個の置換基で置換されたフェニル)であるか、
WPTM2は、任意に置換された6員ヘテロアリール環(例えば、アミノ基で置換されたピリダジンなど、本明細書に記載されるように置換された)であり、
WPTM3は、任意に置換された5~6員ヘテロアリール(例えば、ピラゾール、ピロール、イミダゾール、オキサゾール、オキサジアゾール、またはトリアゾール)であるか、
WPTM5は、本明細書に記載される任意の態様または実施形態に記載されるとおりである(例えば、WPTM5は、存在しないか、またはピリジン環であり得る)か、あるいは
それらの組み合わせ、のうちの少なくとも1つである。
In any aspect or embodiment described herein, the PTM of the PROTAC of the disclosure is represented by Formula I, wherein:
WPTM1 is an optionally substituted phenyl or pyridyl (e.g., substituted as described herein, e.g., substituted with hydroxy or phosphate substituents with or without additional optional substituents selected as described herein, e.g., phenyl substituted with 0, 1, 2, or 3 substituents selected from hydroxy, halogen, alkoxy, alkyl, haloalkyl, amino, alkylamino, cyano, or combinations thereof);
WPTM2 is an optionally substituted 6-membered heteroaryl ring (e.g., substituted as described herein, such as pyridazine substituted with an amino group);
WPTM3 is an optionally substituted 5-6 membered heteroaryl (e.g., pyrazole, pyrrole, imidazole, oxazole, oxadiazole, or triazole);
WPTM5 is at least one of: as described in any aspect or embodiment described herein (e.g., WPTM5 can be absent or a pyridine ring); or a combination thereof.
本明細書に記載される任意の態様または実施形態、例えば、式IのPTMを含む実施形態では、WPTM3は、ピラゾールまたは6~8員ヘテロシクリル(例えば、ピペラジンまたはジアザビシクロオクタン)である。 In any aspect or embodiment described herein, including embodiments that include a PTM of formula I, WPTM3 is pyrazole or a 6-8 membered heterocyclyl (e.g., piperazine or diazabicyclooctane).
本明細書に記載される任意の態様または実施形態では、本開示のPTMは、以下によって表される化学構造を有し、
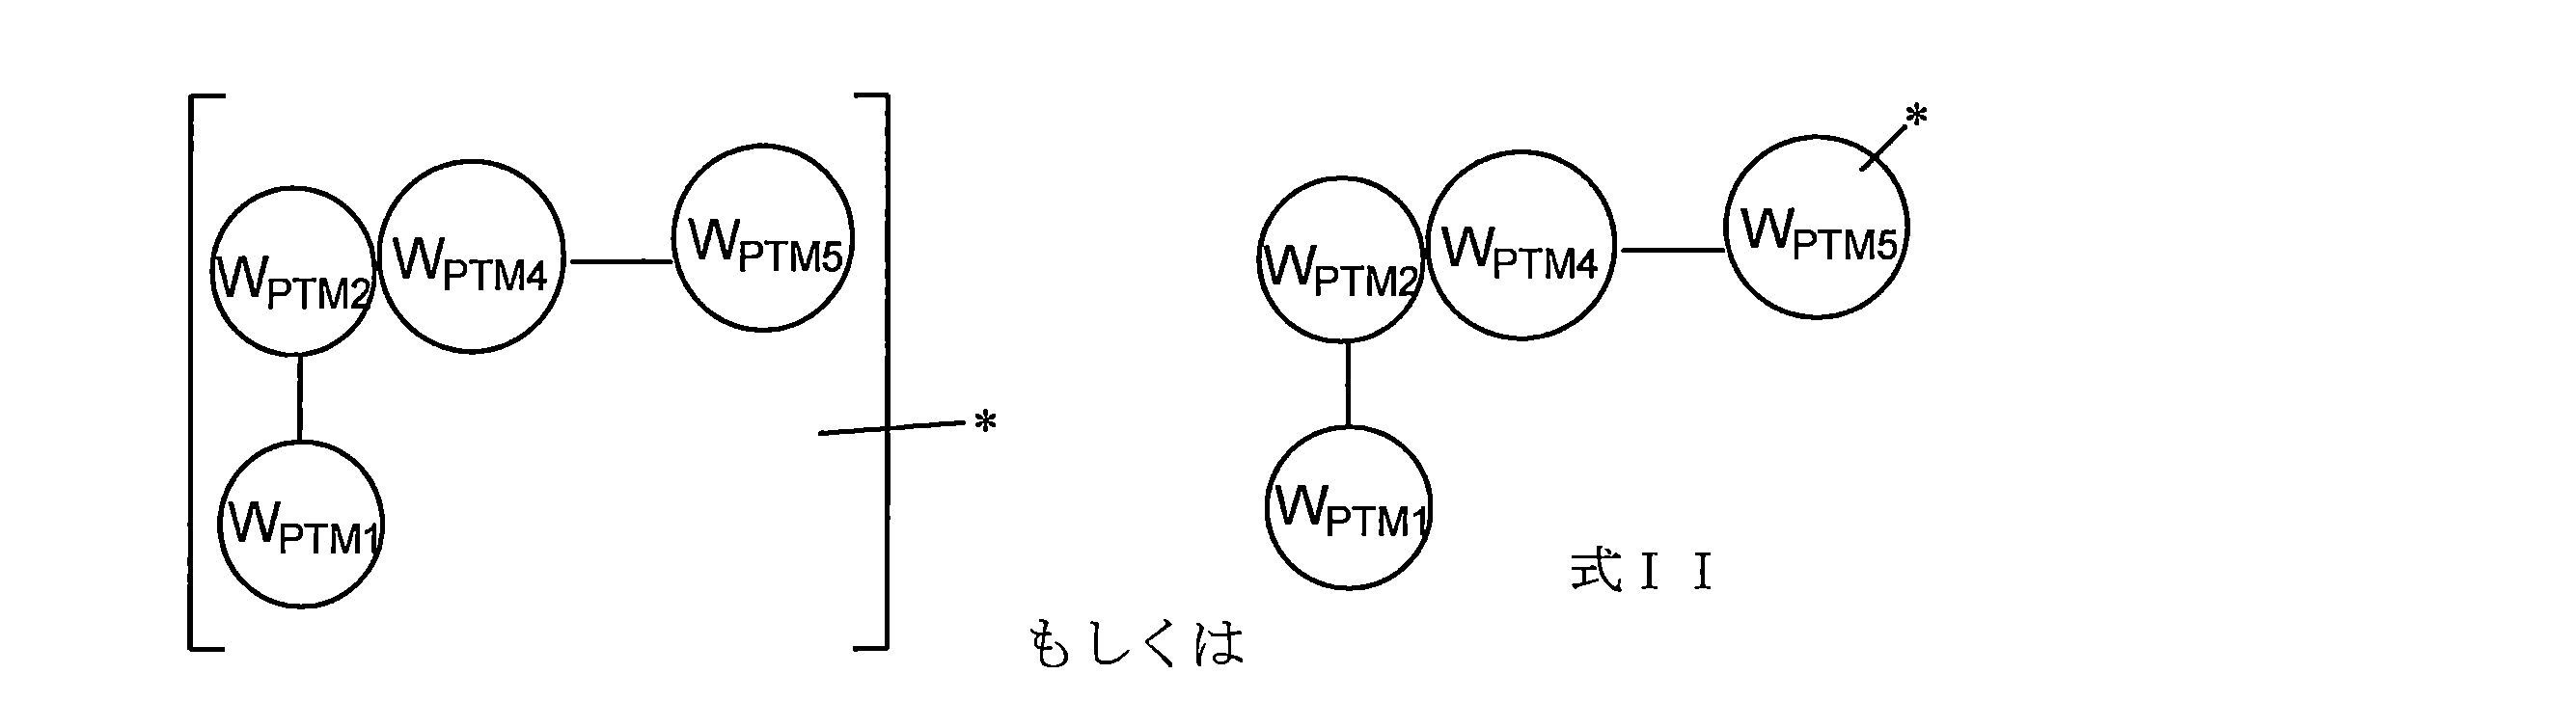
WPTM1、WPTM2、およびWPTM5は、本明細書に記載される任意の他の態様または実施形態に記載されるとおり(例えば、WPTM5は、WPTM4がL(linker)またはULMに直接連結し得るため、存在してもしなくてもよい)であり、
WPTM4は、WPTM2環で縮合された、任意に置換された3~7シクロアルキルまたはヘテロシクリル(例えば、任意に置換された5~7シクロアルキルもしくはヘテロシクリル、またはヒドロキシ、ハロゲン、アルコキシ、アルキル、ハロアルキル、アミノ、アルキルアミノ、およびシアノから選択される0、1、2、または3個の置換基で置換された5~7シクロアルキルまたはヘテロシクリル)であり、

WPTM1, WPTM2, and WPTM5 are as described in any other aspect or embodiment described herein (e.g., WPTM5 may be present or absent because WPTM4 may directly link to L (linker) or ULM);
WPTM4 is an optionally substituted 3-7 cycloalkyl or heterocyclyl (e.g., an optionally substituted 5-7 cycloalkyl or heterocyclyl, or a 5-7 cycloalkyl or heterocyclyl substituted with 0, 1, 2, or 3 substituents selected from hydroxy, halogen, alkoxy, alkyl, haloalkyl, amino, alkylamino, and cyano) fused to a WPTM2 ring;
本明細書に記載される任意の態様または実施形態では、本開示のPTMは、式IIによって表され、式中、WPTM1、WPTM2、およびWPTM5は、本明細書に記載される態様または実施形態のいずれかに記載されるとおりであり、WPTM4は、ピペラジン環である。例えば、本明細書に記載される任意の態様または実施形態では、式IIのWPTM2およびWPTM4は一緒になって、以下に示されるジヒドロピラジノ[2,3-e]ピリダジンを構成する。

本明細書に記載される任意の態様または実施形態では、本開示のPTMは、以下によって表される化学構造を有し、
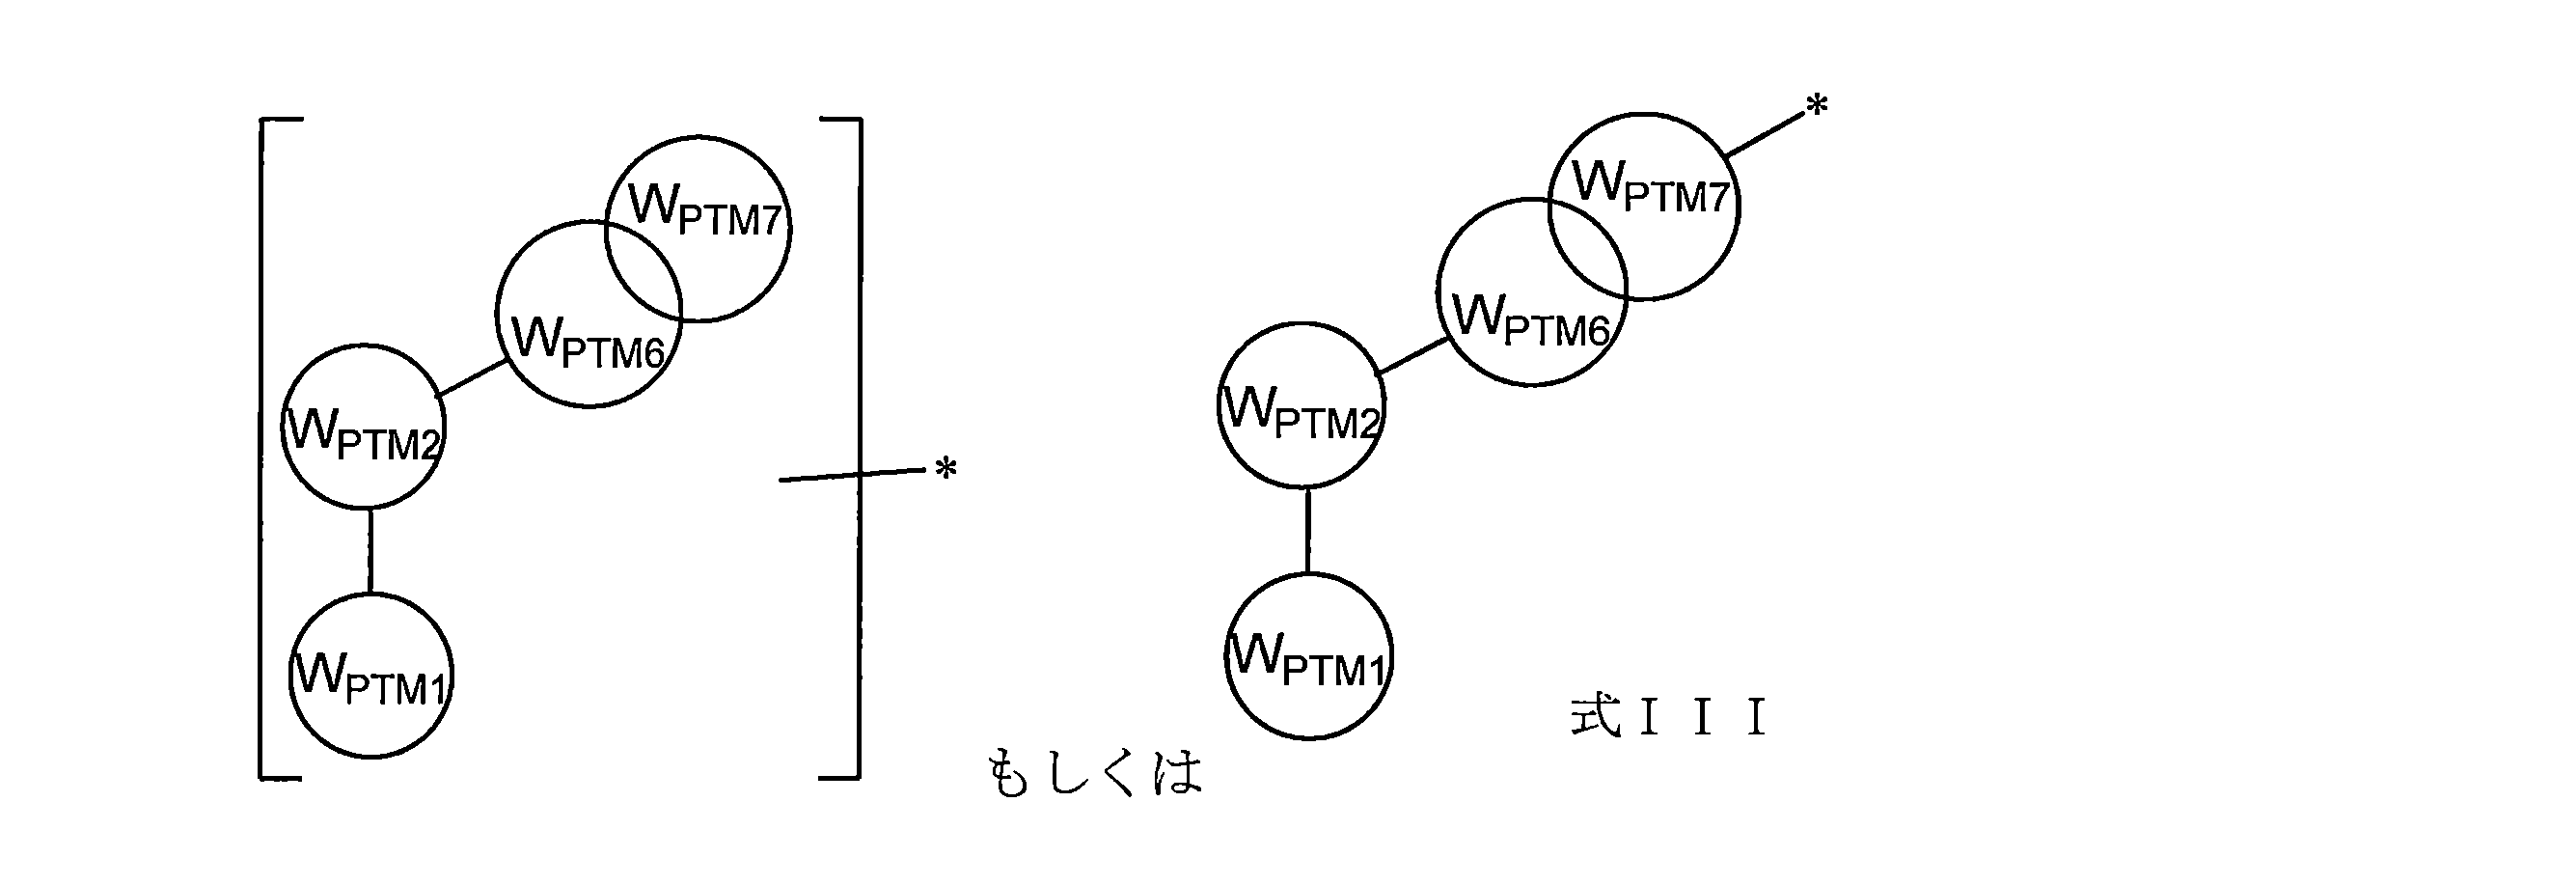
WPTM1およびWPTM2は、本明細書に記載される任意の態様または実施形態に記載されるとおりであり、
WPTM6およびWPTM7は、独立して、任意に4~7シクロアルキルまたはヘテロシクリル(例えば、各々が独立して、ヒドロキシ、ハロゲン、アルコキシ、アルキル、ハロアルキル、アミノ、アルキルアミノ、およびシアノから選択される0、1、または2個の置換基で置換された4~7シクロアルキルまたはヘテロシクリルである)であり、WPTM6およびWPTM7の環は、スピロ連結を介して縮合または結合され、
WPTM1 and WPTM2 are as described in any aspect or embodiment described herein;
WPTM6 and WPTM7 are independently optionally 4-7 cycloalkyl or heterocyclyl (e.g., 4-7 cycloalkyl or heterocyclyl each independently substituted with 0, 1, or 2 substituents selected from hydroxy, halogen, alkoxy, alkyl, haloalkyl, amino, alkylamino, and cyano), and the rings of WPTM6 and WPTM7 are fused or joined via a spiro linkage;
本明細書に記載される任意の態様または実施形態では、本開示のPTMは、式IIIによって表される化学構造を有し、式中、WPTM1およびWPTM2は、各々独立して、本明細書に記載される任意の態様または実施形態に記載されるように選択され(例えば、WPTM1は、本明細書に記載される追加の任意の置換基を有するかもしくは有さないヒドロキシ置換基で置換されたフェニルであり、WPTM2は、アミノ基で置換されたピリダジンである)、WPTM6およびWPTM7は、スピロ環状環系、例えば、以下から選択されるスピロ環状環である。

本明細書に記載される任意の態様または実施形態では、本開示のPTMは、以下によって表され、
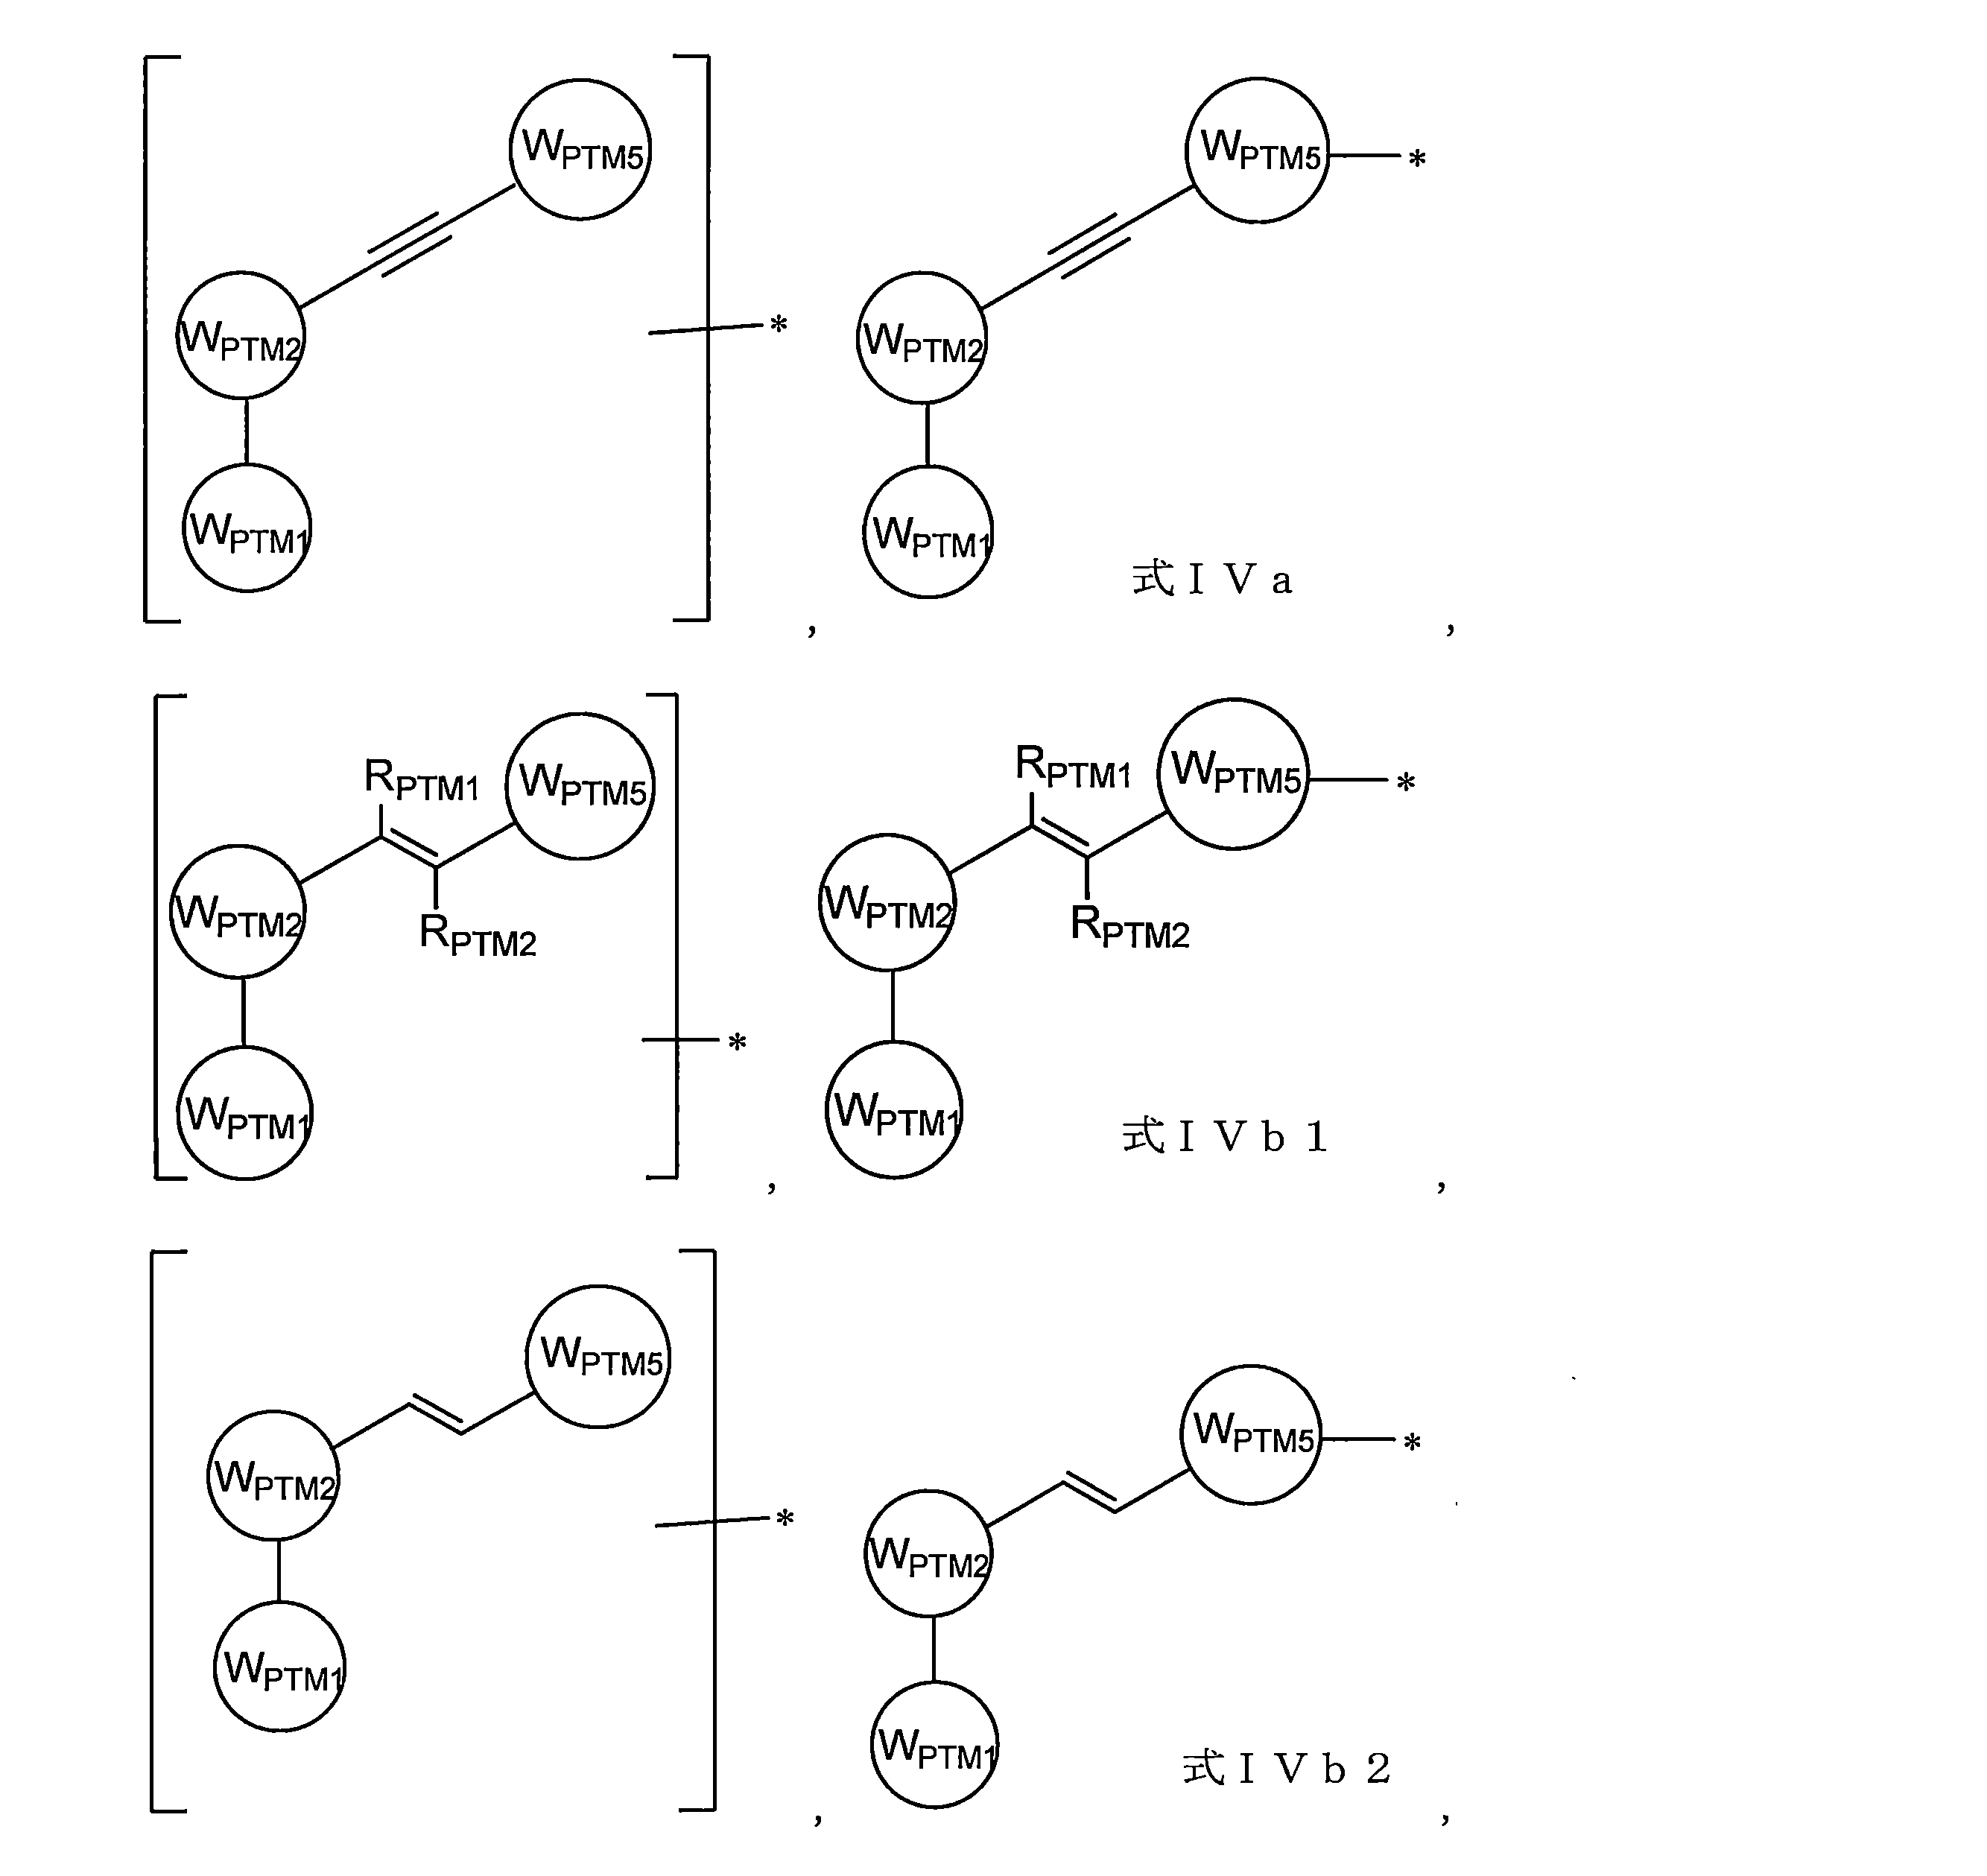
WPTM1、WPTM2、およびWPTM5は、本明細書に記載される任意の他の態様または実施形態に記載されるとおりであり、
RPTM1およびRPTM2は、個々に、H、ハロ、HO、C1-C3アルキル、C1-C3ハロアルキ、またはC1-C3アルコキシであり、

WPTM1, WPTM2, and WPTM5 are as described in any other aspect or embodiment described herein;
RPTM1 and RPTM2 are individually H, halo, HO, C1-C3 alkyl, C1-C3 haloalkoxy, or C1-C3 alkoxy;
本明細書に記載される任意の態様または実施形態では、本開示のPTMは、式IVで表される化学構造を有し、式中、
WPTM1は、本明細書に記載される追加の任意の置換基を有するかまたは有しない、ヒドロキシまたはホスフェート置換基で置換された、フェニルであるか、
WPTM2は、アミノ基で置換されたピリダジンであり、
WPTM5は、存在しないか、またはピラゾール環もしくはピリジン環であるか、あるいは
それらの組み合わせ、のうちの少なくとも1つである。
In any aspect or embodiment described herein, the PTM of the present disclosure has a chemical structure represented by Formula IV, wherein:
WPTM1 is a phenyl substituted with a hydroxy or phosphate substituent, with or without any additional substituents described herein;
WPTM2 is an amino-substituted pyridazine;
WPTM5 is at least one of: absent, a pyrazole ring, or a pyridine ring, or a combination thereof.
本明細書に記載される任意の態様または実施形態では、本開示のPTMは、以下によって表されるか、

WPTM3は、不在であるか、または任意に置換された5~6員ヘテロアリール、任意に置換された4~9シクロアルキルもしくはヘテロシクリル環、任意に置換された架橋ビシクロアルキル、および架橋ビヘテロシクリル環であり、
WPTM5は、任意に置換された5~6員ヘテロアリールまたはアリール、例えば、ピリジン、またはピリダジンである。
In any aspect or embodiment described herein, the PTM of the present disclosure is represented by:
WPTM3 is absent or an optionally substituted 5-6 membered heteroaryl, an optionally substituted 4-9 membered cycloalkyl or heterocyclyl ring, an optionally substituted bridged bicycloalkyl, and a bridged biheterocyclyl ring;
WPTM5 is an optionally substituted 5-6 membered heteroaryl or aryl, for example, pyridine, or pyridazine.
本明細書に記載される任意の態様または実施形態では、本開示のPTMは、以下によって表されるか、
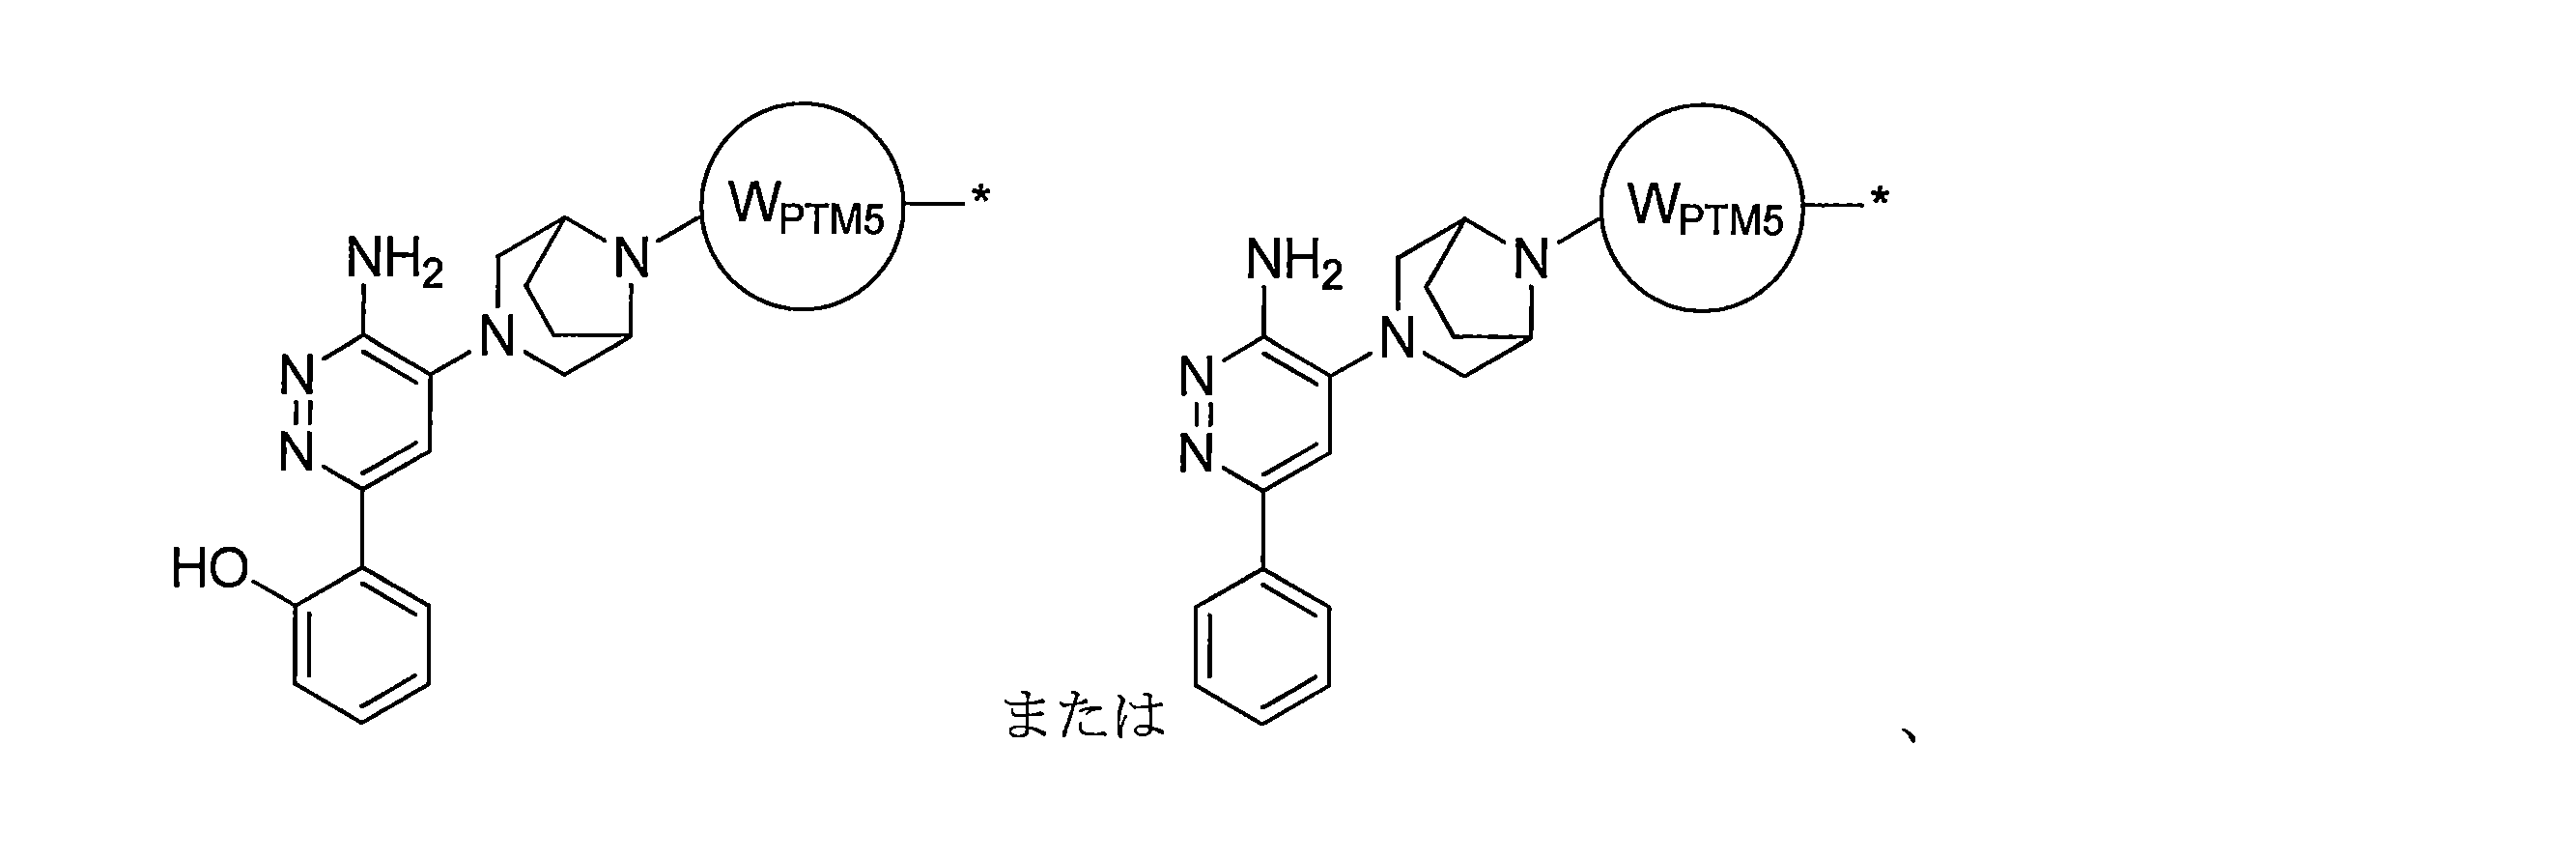
WPTM5は、フェニル、ピリジン、ピリミジン、またはピラジンである。
In any aspect or embodiment described herein, the PTM of the disclosure is represented by:
WPTM5 is phenyl, pyridine, pyrimidine, or pyrazine.
本明細書に記載される任意の態様または実施形態では、本開示のPTMは、以下によって表されるか、

本明細書に記載される任意の態様または実施形態では、本開示のPTMは、以下によって表されるか、

式中、
WPTM3は、任意に置換された5~6員ヘテロアリール、任意に置換された4~9シクロアルキルもしくはヘテロシクリル環、任意に置換された架橋ビシクロアルキルおよび架橋ビヘテロシクリル環であり、
WPTM5は、任意に置換された5~6員ヘテロアリールまたはアリール、例えば、ピリジン、またはピリダジンであり、
Rvは、独立して、ヒドロキシ、ハロゲン、アルコキシ、アルキル、ハロアルキル、ホスフェート、アミノ、アルキルアミノ、シアノ、またはそれらの組み合わせから選択される、0、1、2、または3個の置換基である。
In any aspect or embodiment described herein, the PTM of the disclosure is represented by:
In the formula,
WPTM3 is an optionally substituted 5-6 membered heteroaryl, an optionally substituted 4-9 membered cycloalkyl or heterocyclyl ring, an optionally substituted bridged bicycloalkyl and a bridged biheterocyclyl ring;
WPTM5 is an optionally substituted 5-6 membered heteroaryl or aryl, e.g., pyridine, or pyridazine;
Rv is 0, 1, 2, or 3 substituents independently selected from hydroxy, halogen, alkoxy, alkyl, haloalkyl, phosphate, amino, alkylamino, cyano, or combinations thereof.
ある特定の実施形態では、ヒドロキシル基は、ホスフェート基(すなわち、ホスホエステル基)で修飾される。 In certain embodiments, the hydroxyl groups are modified with phosphate groups (i.e., phosphoester groups).
本明細書に記載される任意の態様または実施形態では、本開示のPTMは、以下によって表される化学構造を有し、

WPTM3は、存在しないか、またはWPTM2環で縮合された、任意に置換された5~7シクロアルキルもしくはヘテロシクリル(例えば、ヒドロキシ、ハロゲン、アルコキシ、アルキル、ハロアルキル、アミノ、アルキルアミノ、およびシアノから選択される0、1、2、もしくは3個の置換基で置換された5~7シクロアルキルもしくはヘテロシクリル)であり、
WPTM4は、任意に置換された3~7員アリールもしくはヘテロアリール環(例えば、任意に置換された5~7シクロアルキルもしくはヘテロシクリル、またはヒドロキシ、ハロゲン、アルコキシ、アルキル、ハロアルキル、アミノ、アルキルアミノ、およびシアノから選択される0、1、2、もしくは3個の置換基で置換された3~7もしくは5~6員アリールもしくはヘテロアリール)、または任意に置換された4~9シクロアルキルもしくはヘテロシクリル、例えば、任意に置換された架橋ビシクロアルキルおよび架橋ビヘテロシクリル環(例えば、ヒドロキシ、ハロゲン、アルコキシ、アルキル、ハロアルキル、アミノ、アルキルアミノ、およびシアノから選択される0、1、もしくは2個の置換基で置換された4~9シクロアルキルもしくはヘテロシクリル)であり、
WPTM5は、存在しない(例えば、WPTM3が、L(リンカー)またはULMに直接連結されている)か、または任意に置換されたアルキル、任意に置換された5~6員シクロアルキル、複素環、アリール、もしくはヘテロアリール環(例えば、ヒドロキシ、ハロゲン、アルコキシ、アルキル、ハロアルキル、アミノ、アルキルアミノ、およびシアノから選択される0、1、または2個の置換基で置換された、5~6員シクロアルキル、複素環、アリール、またはヘテロアリール)、例えば、任意に置換されたピラゾール環、またはピリジン環であり、

WPTM3 is absent or is an optionally substituted 5-7 cycloalkyl or heterocyclyl (e.g., 5-7 cycloalkyl or heterocyclyl substituted with 0, 1, 2, or 3 substituents selected from hydroxy, halogen, alkoxy, alkyl, haloalkyl, amino, alkylamino, and cyano) fused to a WPTM2 ring;
WPTM4 is an optionally substituted 3-7 membered aryl or heteroaryl ring (e.g., an optionally substituted 5-7 cycloalkyl or heterocyclyl, or a 3-7 or 5-6 membered aryl or heteroaryl substituted with 0, 1, 2, or 3 substituents selected from hydroxy, halogen, alkoxy, alkyl, haloalkyl, amino, alkylamino, and cyano), or an optionally substituted 4-9 cycloalkyl or heterocyclyl, such as optionally substituted bridged bicycloalkyl and bridged biheterocyclyl rings (e.g., a 4-9 cycloalkyl or heterocyclyl substituted with 0, 1, or 2 substituents selected from hydroxy, halogen, alkoxy, alkyl, haloalkyl, amino, alkylamino, and cyano);
WPTM5 is absent (e.g., WPTM3 is directly linked to L (linker) or ULM) or is an optionally substituted alkyl, an optionally substituted 5-6 membered cycloalkyl, heterocycle, aryl, or heteroaryl ring (e.g., a 5-6 membered cycloalkyl, heterocycle, aryl, or heteroaryl substituted with 0, 1, or 2 substituents selected from hydroxy, halogen, alkoxy, alkyl, haloalkyl, amino, alkylamino, and cyano), e.g., an optionally substituted pyrazole ring, or a pyridine ring;
本明細書に記載される任意の態様または実施形態では、本開示のPTMは、以下によって表される化学構造を有し、

WPTM1、WPTM2、WPTM3、およびWPTM4は、本明細書に記載される任意の他の態様または実施形態に記載されるとおりであり、
LPTMは、C1-C4アルキルまたはC1-C3アルコキシで任意に置換されたC1-C6アルキルであり、

WPTM1, WPTM2, WPTM3, and WPTM4 are as described in any other aspect or embodiment described herein;
LPTM is C1-C6 alkyl optionally substituted with C1-C4 alkyl or C1-C3 alkoxy;
本明細書に記載される任意の態様または実施形態では、PTMは、
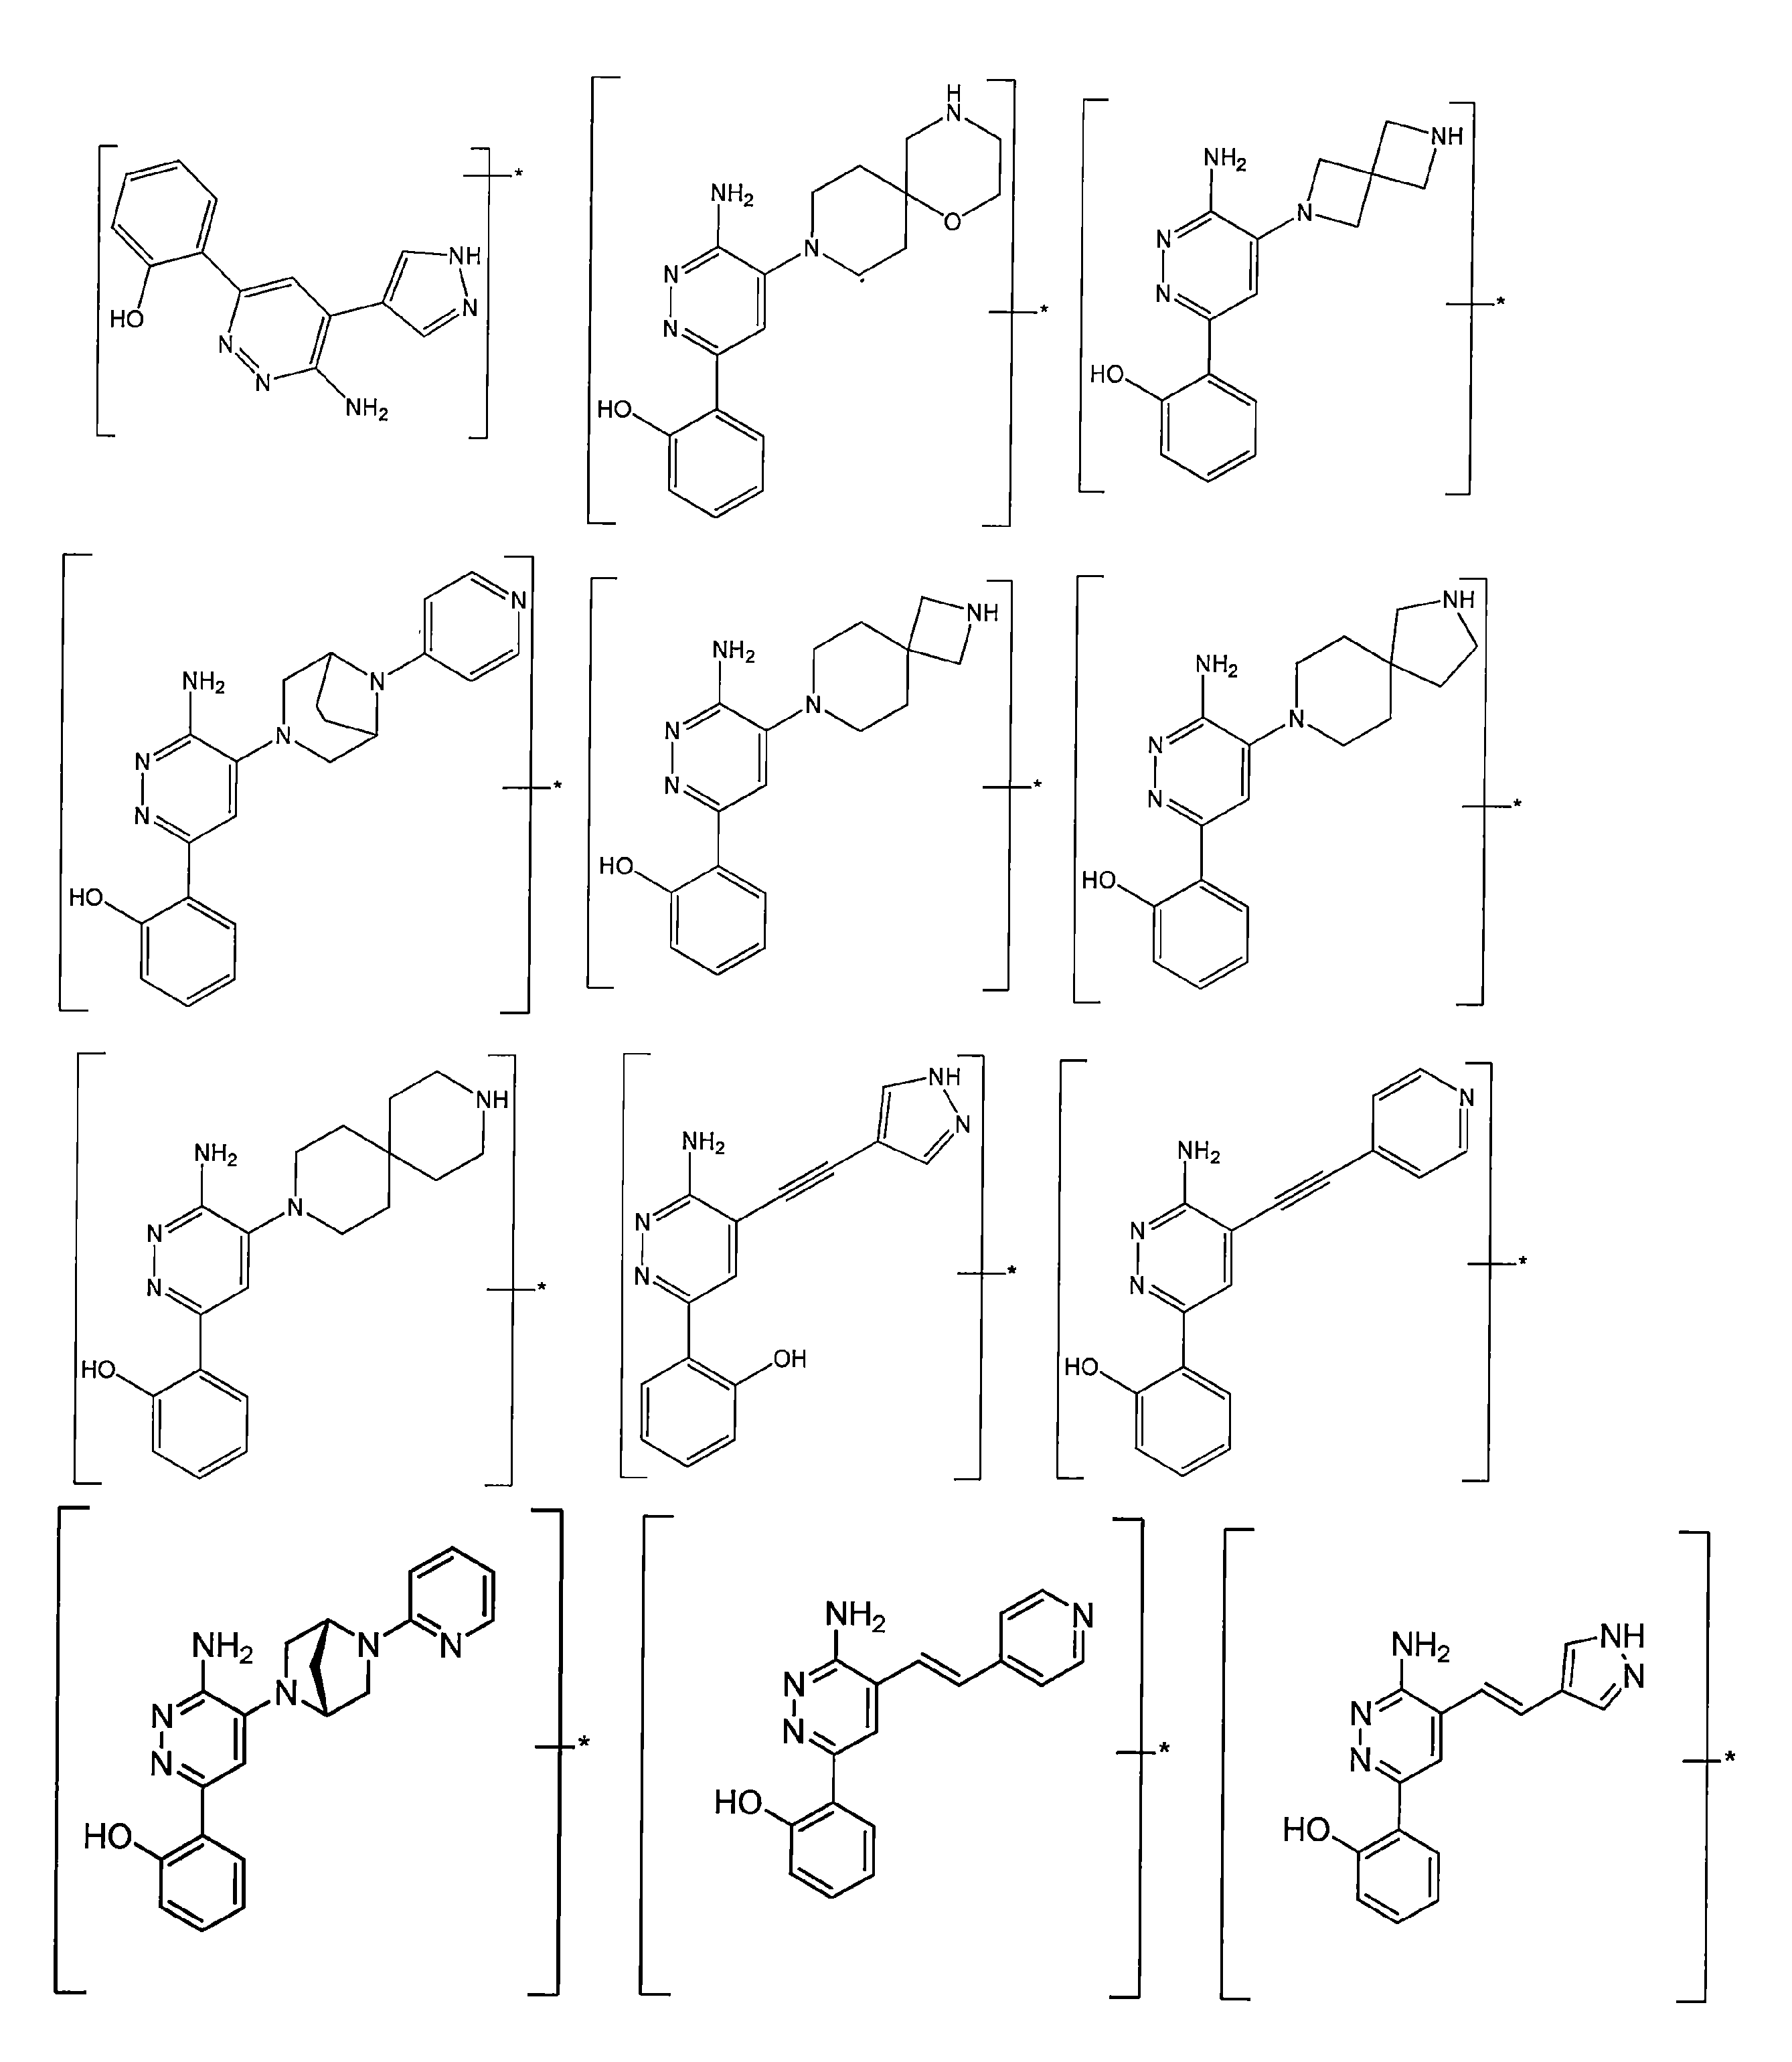



本明細書に記載される任意の態様または実施形態では、PTMは、

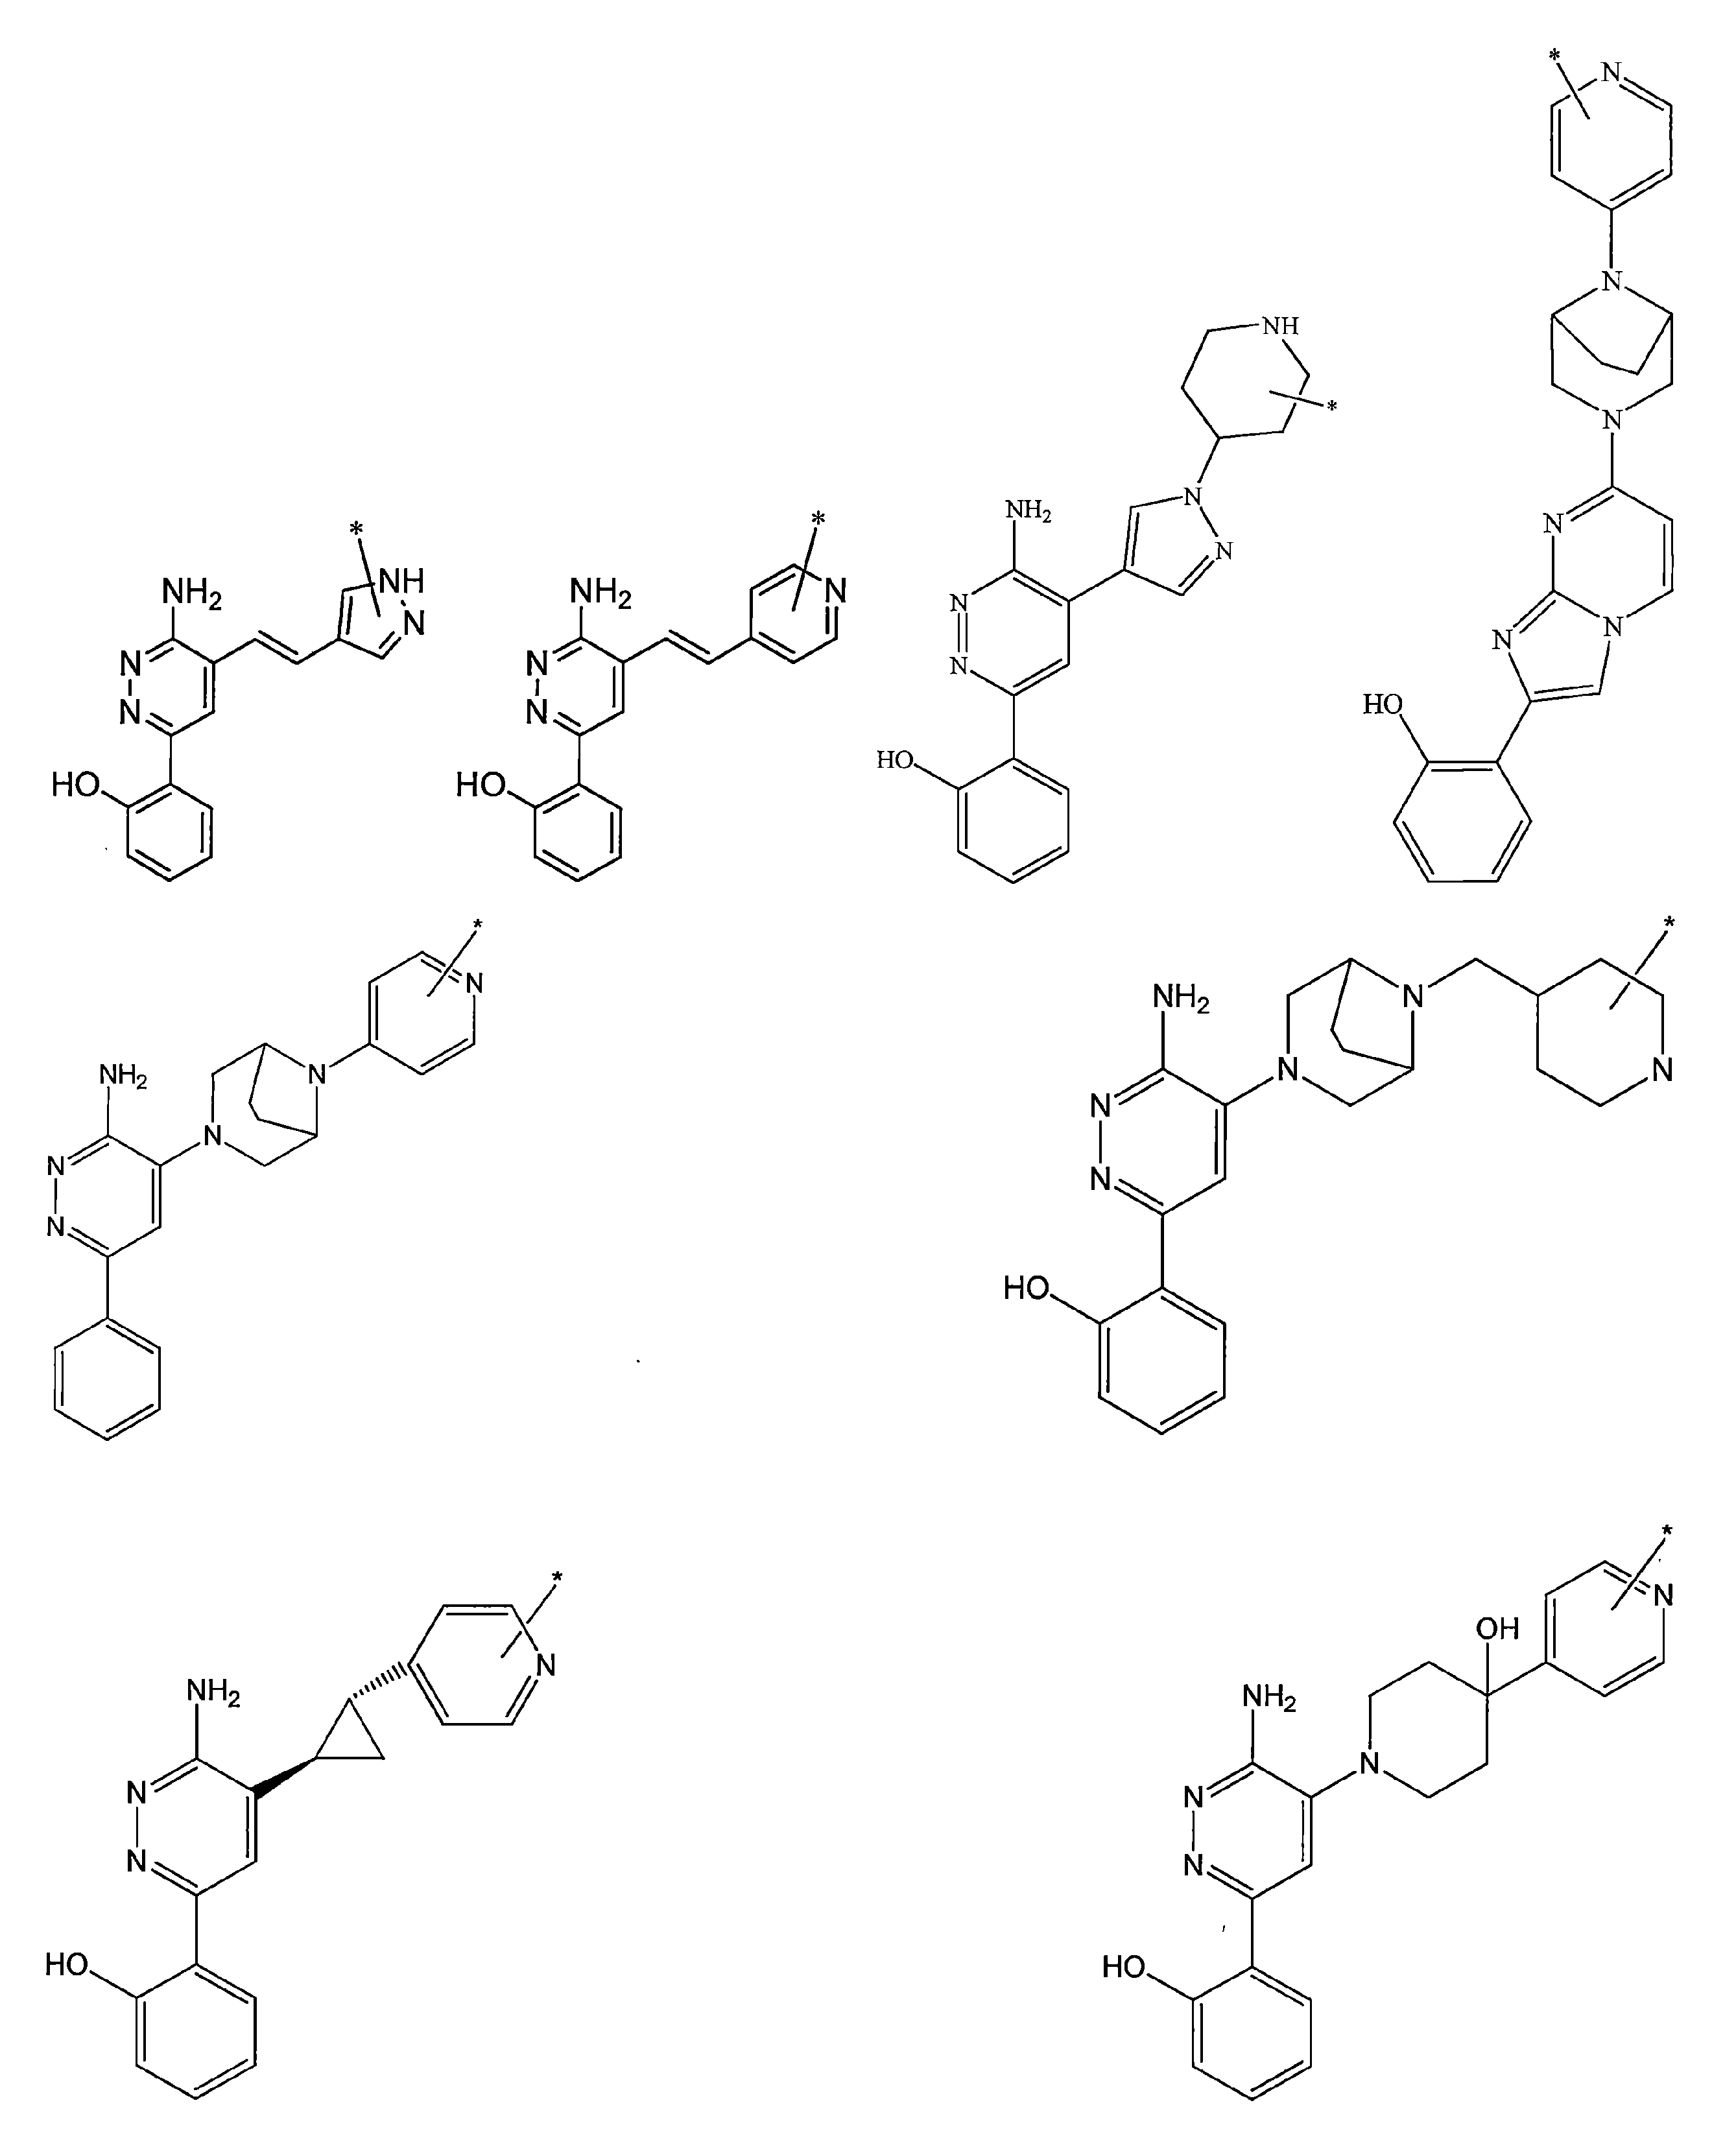
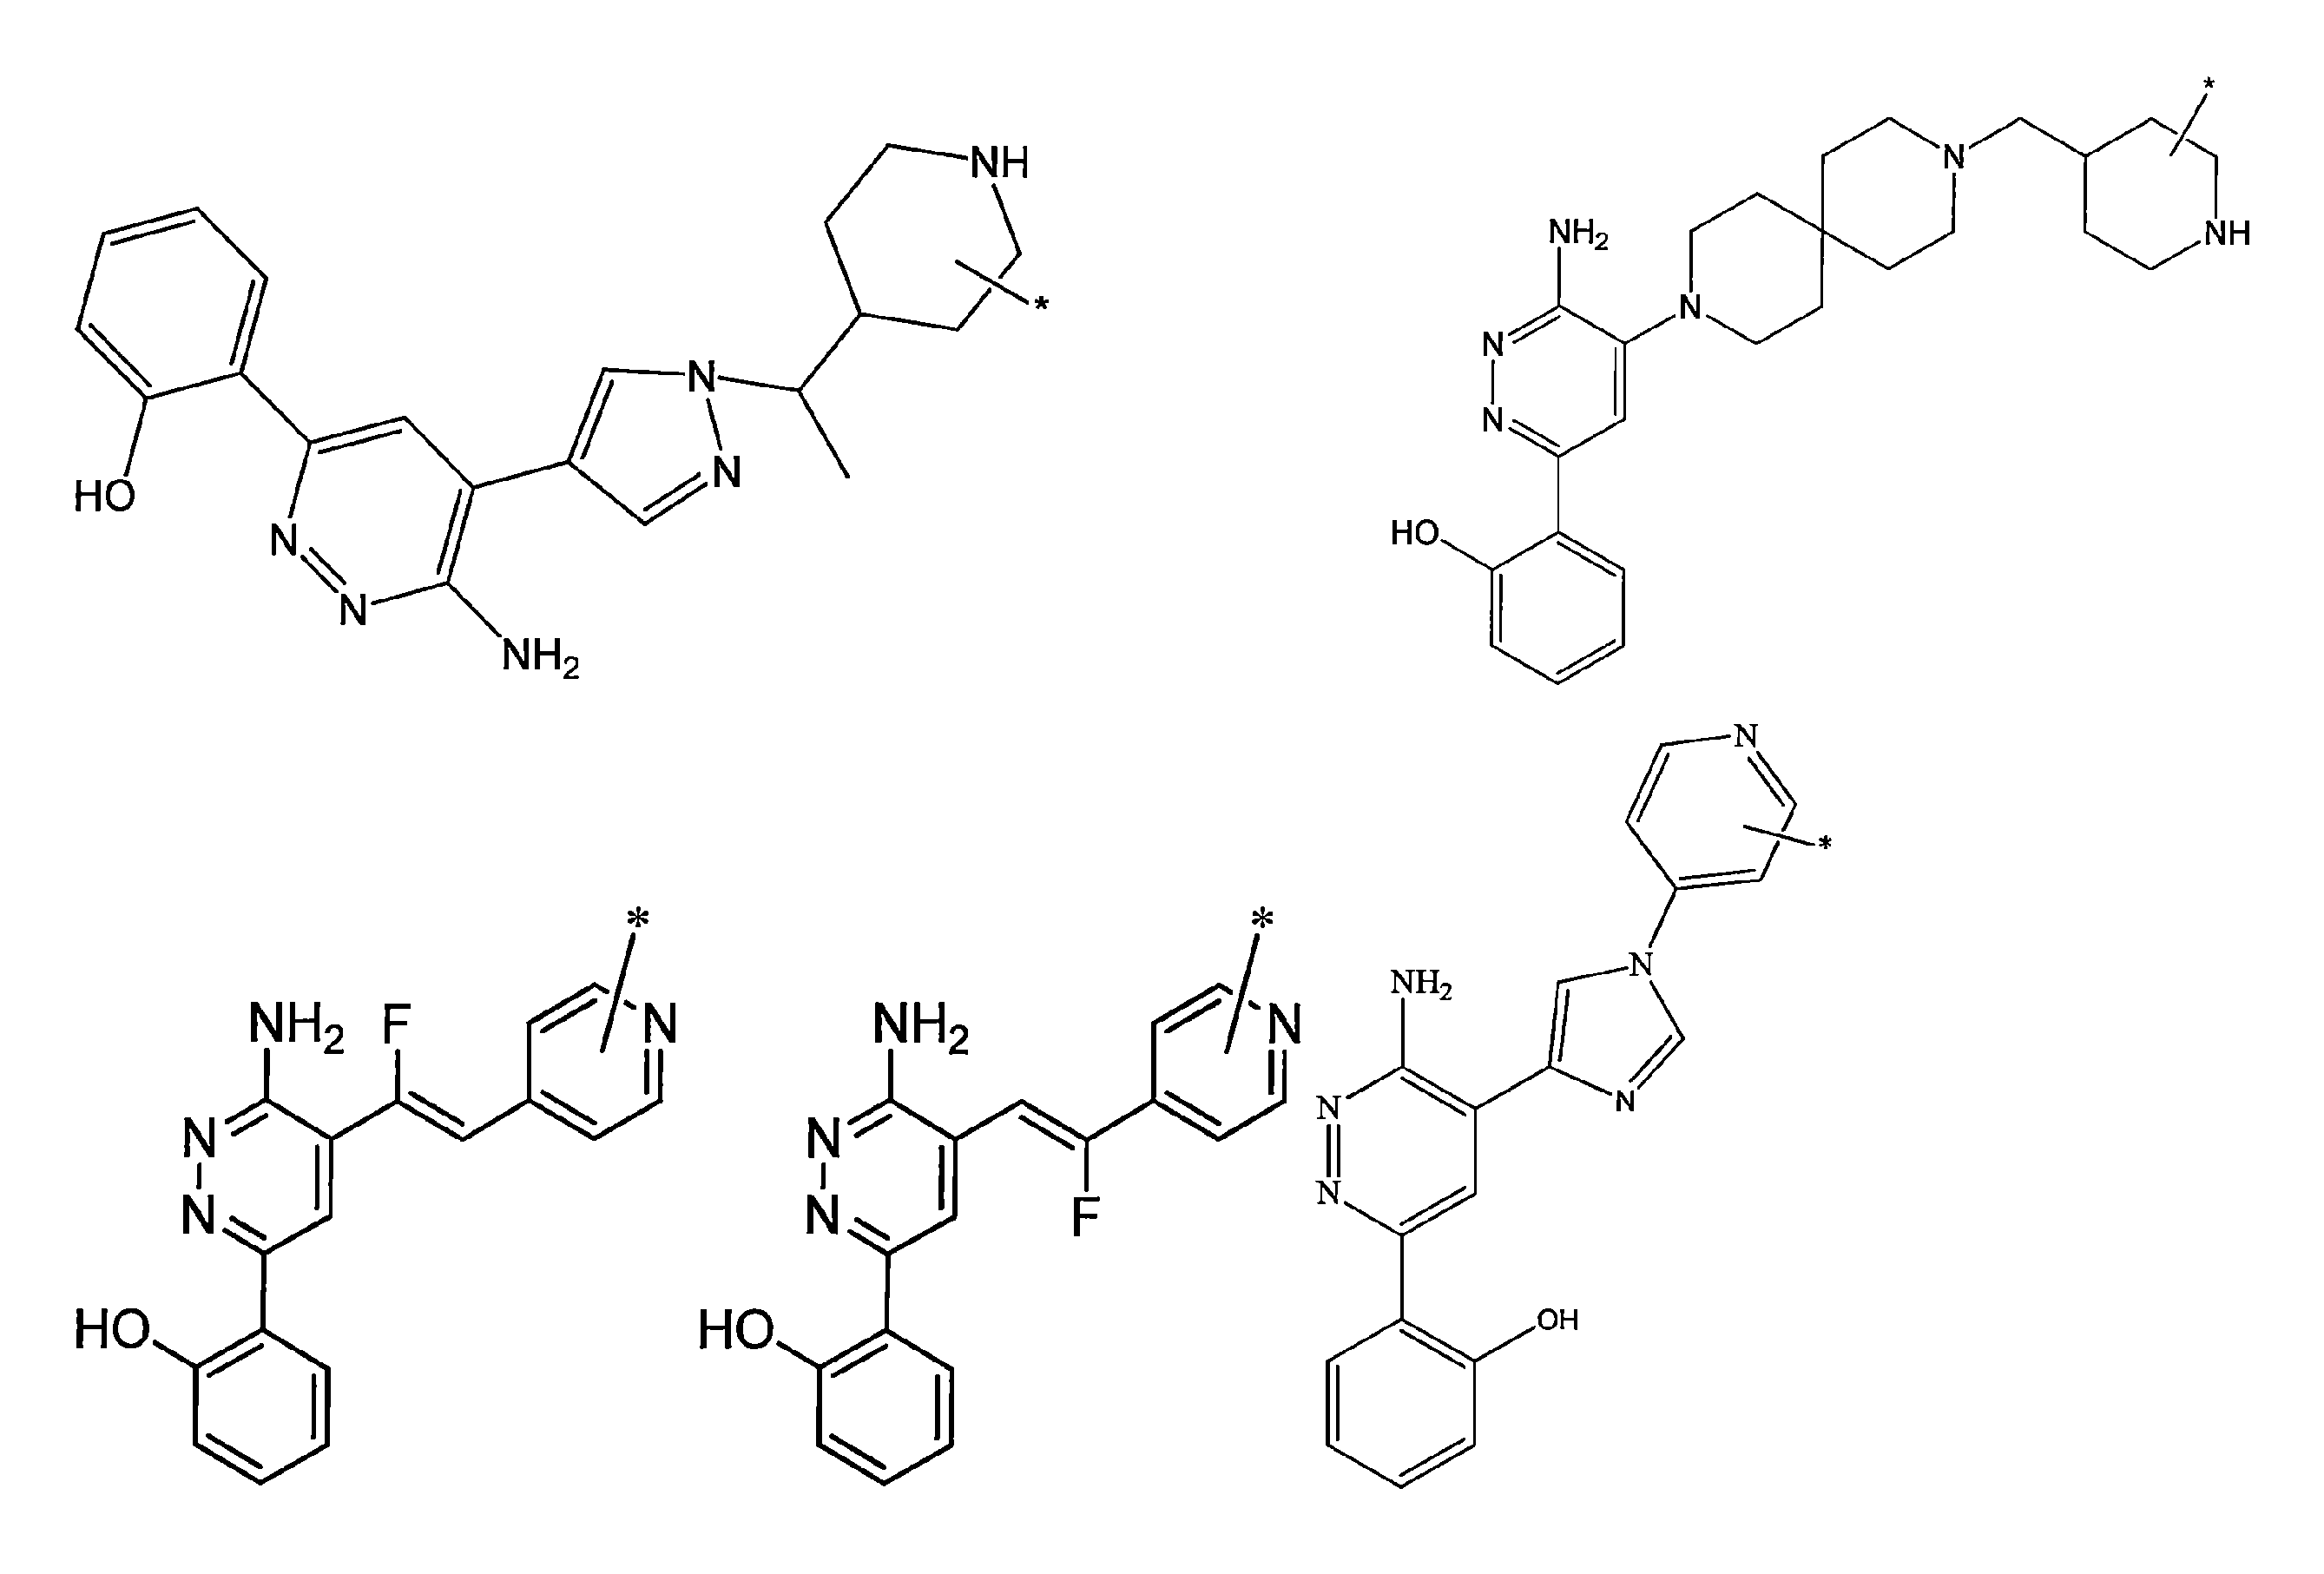

本明細書に記載される任意の態様または実施形態では、PTMは、
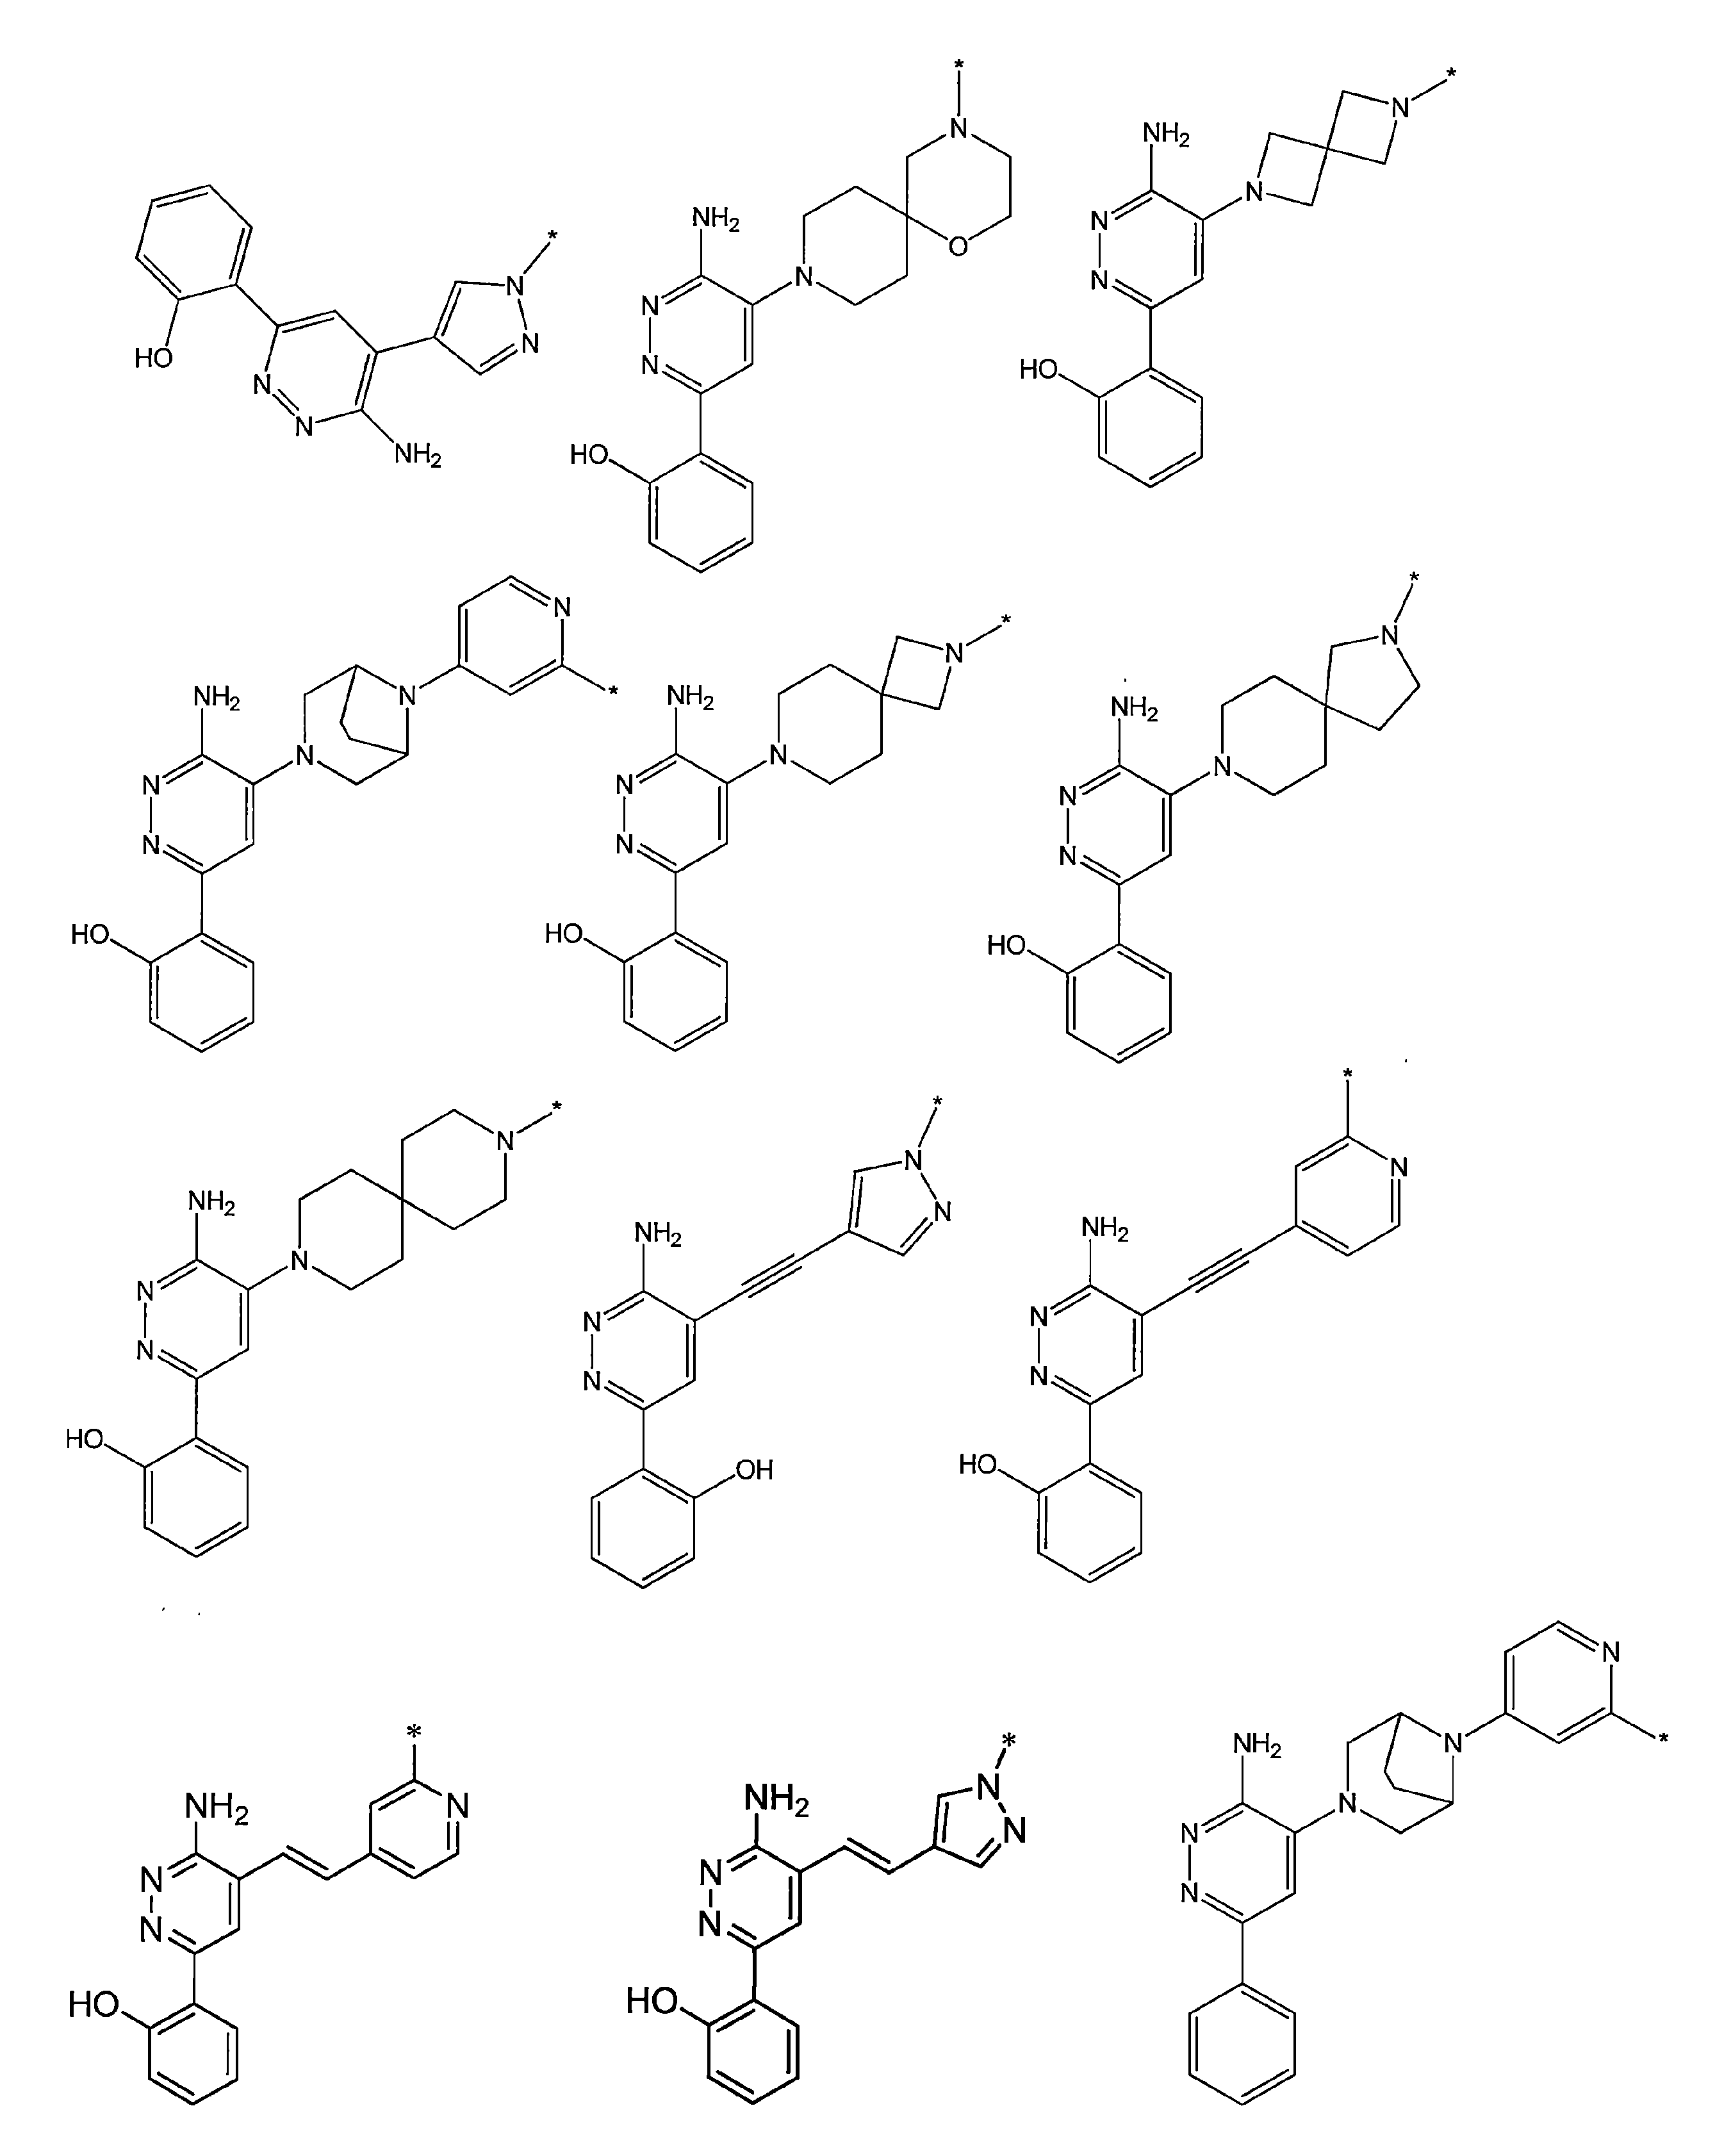
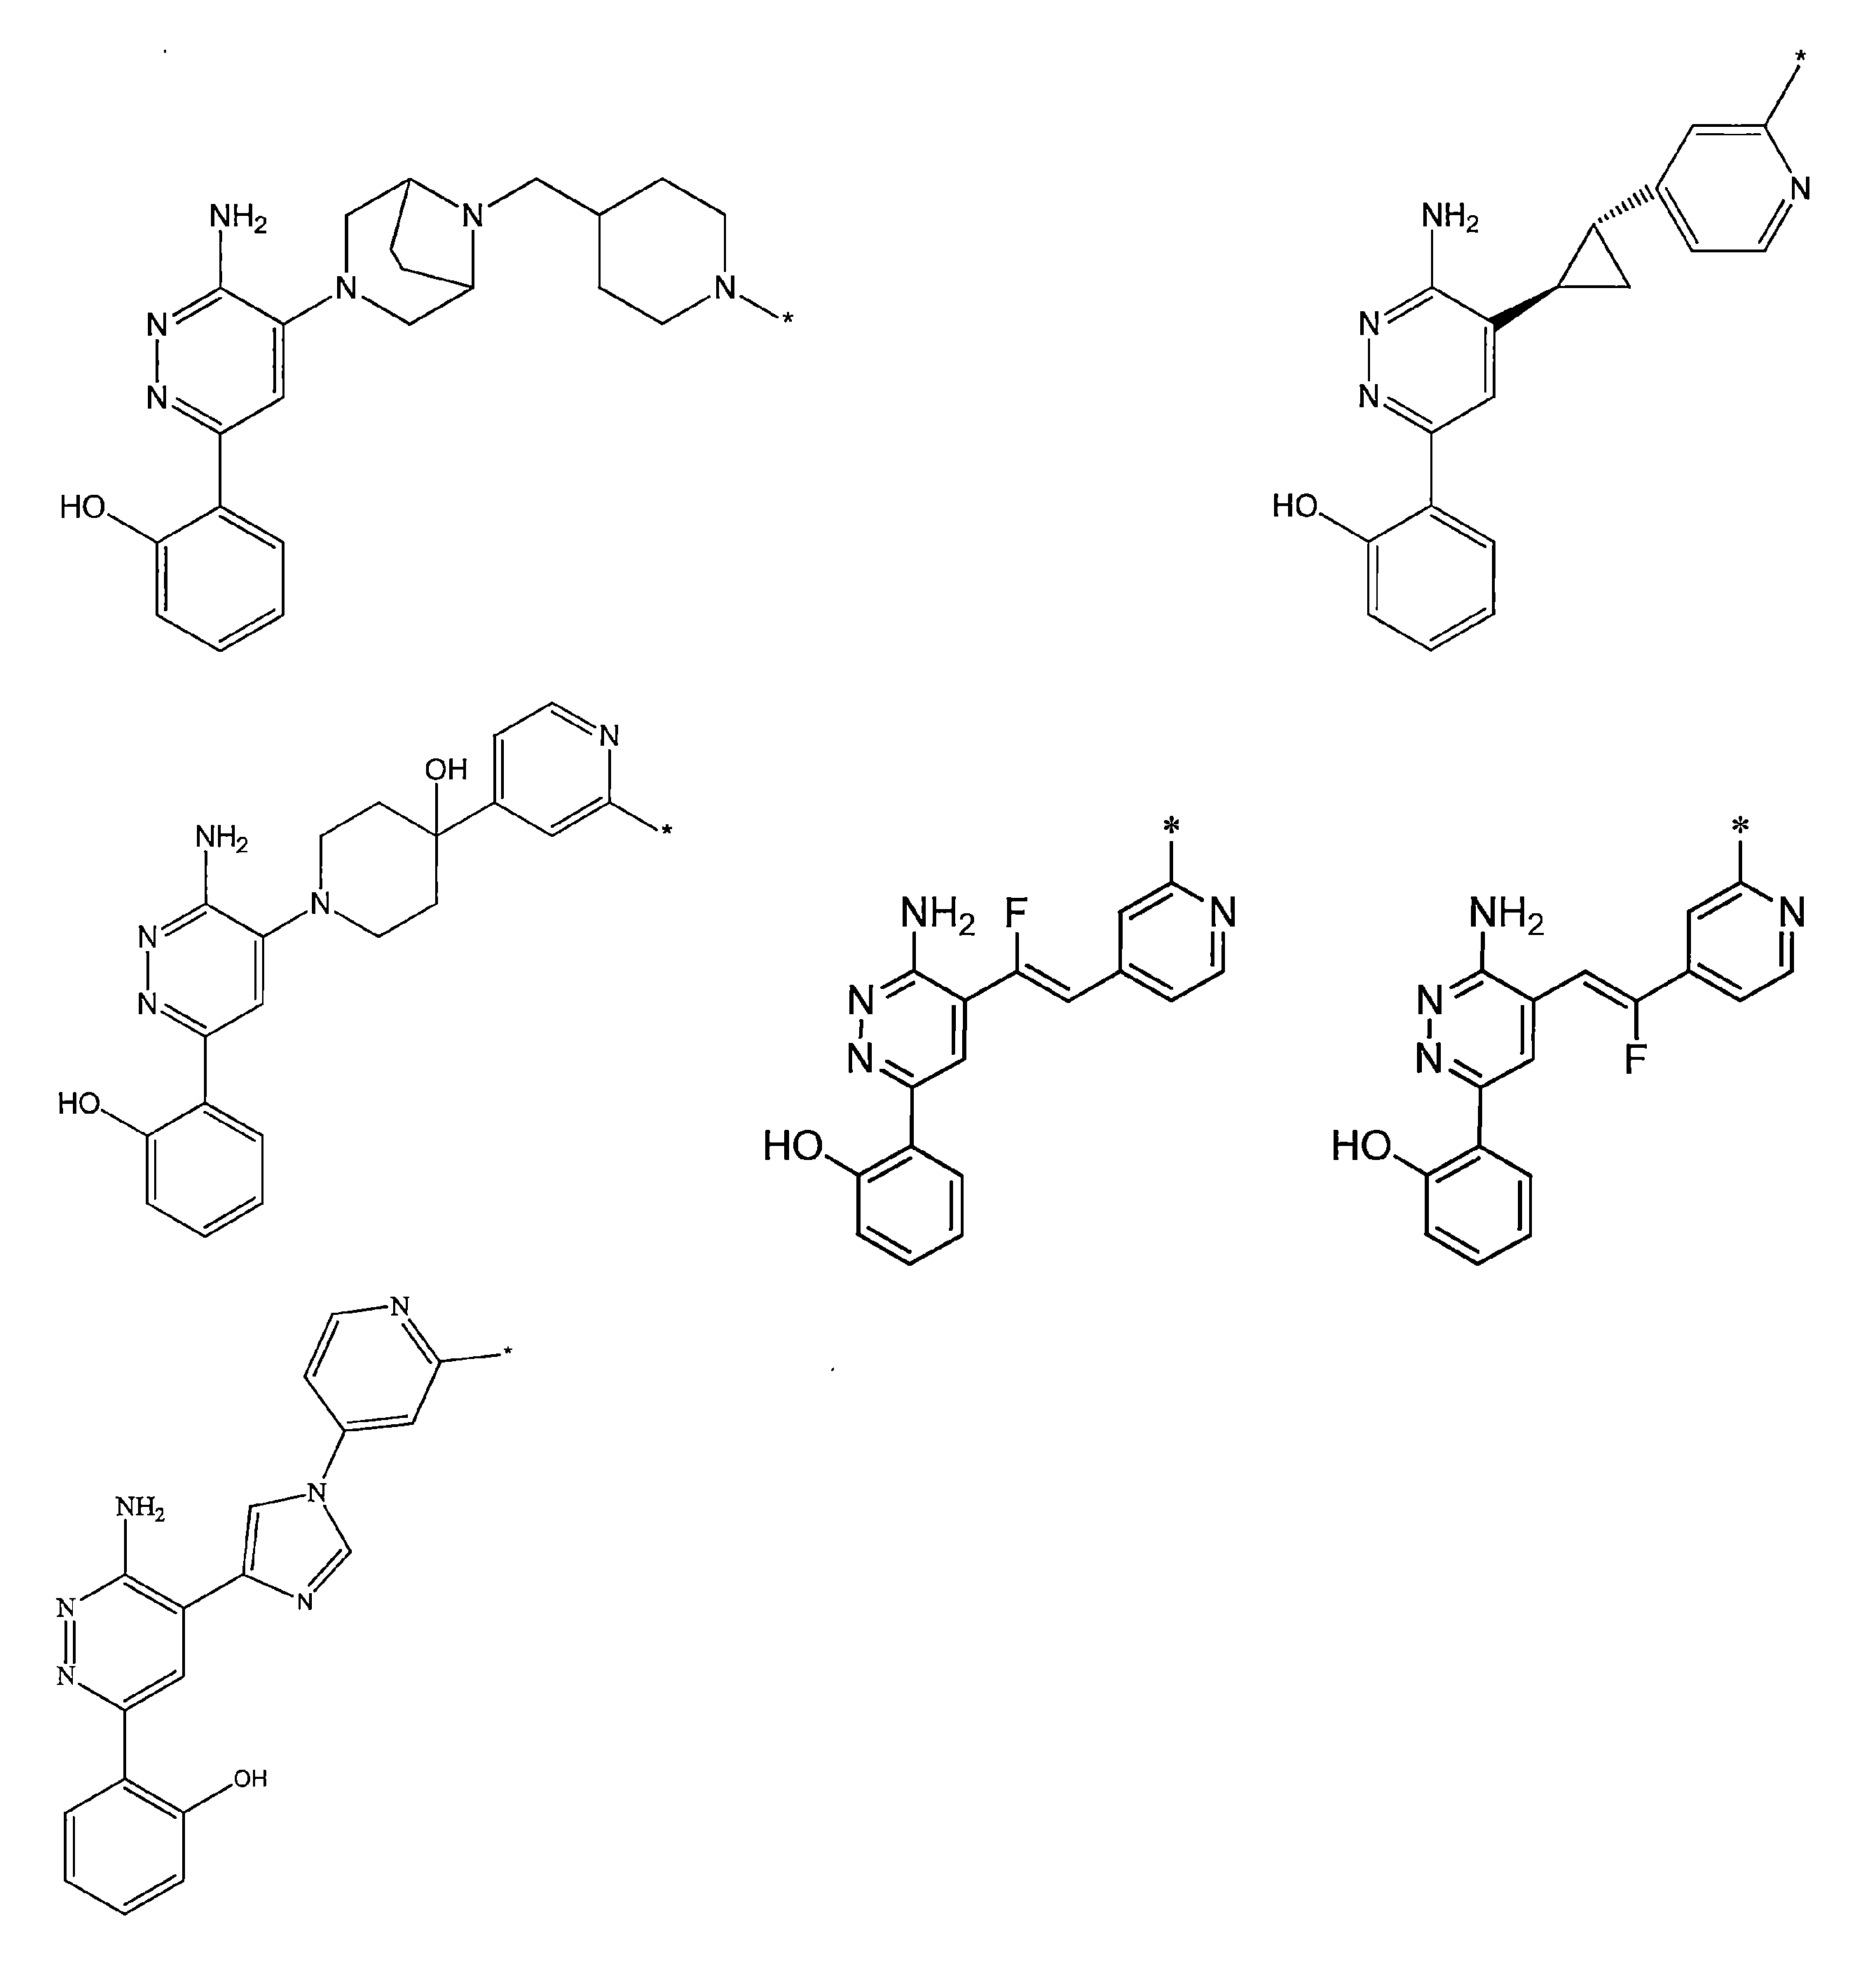

本明細書に記載される組成は、これらのタイプの一連の低分子標的タンパク質結合部分のいくつかを例示する。かかる低分子標的タンパク質結合部分には、これらの組成物の薬学的に許容可能な塩、鏡像異性体、溶媒和物、および多形体、ならびに対象のタンパク質を標的としうる他の低分子も含まれる。以下で本明細書に引用される参照文献は、参照によりその全体が本明細書に組み込まれる。 The compositions described herein exemplify some of these types of small molecule target protein binding moieties. Such small molecule target protein binding moieties also include pharma- ceutically acceptable salts, enantiomers, solvates, and polymorphs of these compositions, as well as other small molecules that may target proteins of interest. The references cited herein below are incorporated by reference in their entireties.
治療用組成物
本明細書に記載される少なくとも1つの二官能性化合物の有効量、および本明細書において別途記載されている化合物のうちの1つ以上の有効量の組み合わせを、薬学的に有効な量の担体、添加剤、または賦形剤と併用して含む医薬組成物は、本開示のさらなる態様を表す。
Therapeutic Compositions Pharmaceutical compositions comprising an effective amount of at least one bifunctional compound described herein, and a combination of an effective amount of one or more of the compounds otherwise described herein, in combination with a pharma- ceutical effective amount of a carrier, excipient, or diluent, represent a further aspect of the present disclosure.
本開示には、該当する場合、本明細書に記載される化合物の薬学的に許容可能な塩、特に酸付加塩または塩基付加塩を含む組成物が含まれる。本態様に従い有用な上述の塩基化合物の薬学的に許容可能な酸付加塩を調製するために使用される酸は、非毒性の酸付加塩、すなわち薬理学的に許容可能なアニオンを含有する塩、例えば特に多くの他の酸のなかでも塩酸塩、臭化水素酸塩、ヨウ化水素酸塩、硝酸塩、硫酸塩、重硫酸塩、リン酸塩、過リン酸塩、酢酸塩、乳酸塩、クエン酸塩、過クエン酸塩、酒石酸塩、重酒石酸塩、コハク酸塩、マレイン酸塩、フマル酸塩、グルコン酸塩、サッカラート、安息香酸塩、メタンスルホン酸塩、エタンスルホン酸塩、ベンゼンスルホン酸塩、p-トルエンスルホン酸塩、およびパモ酸塩[すなわち1,1’-メチレン-ビス-(2-ヒドロキシ-3ナフトエ酸)]を形成する酸である。 The present disclosure includes compositions comprising pharma- ceutically acceptable salts, particularly acid or base addition salts, of the compounds described herein, where applicable. The acids used to prepare pharma- ceutically acceptable acid addition salts of the above-mentioned base compounds useful according to the present embodiment are those that form non-toxic acid addition salts, i.e., salts containing pharmacologically acceptable anions, such as, among many others, hydrochloride, hydrobromide, hydroiodide, nitrate, sulfate, bisulfate, phosphate, perphosphate, acetate, lactate, citrate, percitrate, tartrate, bitartrate, succinate, maleate, fumarate, gluconate, saccharate, benzoate, methanesulfonate, ethanesulfonate, benzenesulfonate, p-toluenesulfonate, and pamoate salts [i.e., 1,1'-methylene-bis-(2-hydroxy-3-naphthoic acid)].
薬学的に許容可能な塩基付加塩は、本開示による化合物または誘導体の薬学的に許容可能な塩形態を生成するためにも使用され得る。性質的に酸性である本化合物の薬学的に許容可能な塩基塩を調製するための試薬として使用されうる化学塩基は、かかる化合物と非毒性の塩基塩を形成する塩基である。かかる非毒性の塩基塩としては限定されないが、特に例えばアルカリ金属カチオン(例えば、カリウムおよびナトリウム)およびアルカリ土類金属カチオン(例えば、カルシウム、亜鉛およびマグネシウム)などの薬学的に許容可能なカチオンから誘導された塩基塩、アンモニウムまたは水に可溶性のアミン添加塩、例えばN-メチルグルカミン-(メグルミン)、ならびに低級アルカノールアンモニウム、ならびに他の薬学的に許容可能な有機アミンの塩基塩が挙げられる。 Pharmaceutically acceptable base addition salts may also be used to generate pharma- ceutically acceptable salt forms of the compounds or derivatives according to the present disclosure. Chemical bases that may be used as reagents to prepare pharma- ceutically acceptable base salts of the present compounds that are acidic in nature are those that form non-toxic base salts with such compounds. Such non-toxic base salts include, but are not limited to, base salts derived from pharma- ceutically acceptable cations, such as alkali metal cations (e.g., potassium and sodium) and alkaline earth metal cations (e.g., calcium, zinc and magnesium), ammonium or water-soluble amine addition salts, such as N-methylglucamine-(meglumine), and lower alkanolammonium, as well as other pharma- ceutically acceptable organic amine base salts.
本明細書に記載の化合物は、本開示に従い、経口、非経口または局所経路により、単回用量で、または分割用量で投与されてもよい。活性化合物の投与は、連続投与(静脈内点滴)から1日あたり数回の経口投与(例えば、Q.I.D.)の範囲であり得、投与経路の中でもとりわけ、経口、局所、非経口、筋肉内、静脈内、皮下、経皮(貫通増強剤を含み得る)、口腔、舌下、および座薬投与が挙げられる。腸溶コーティングされた経口錠剤を使用して、経口投与経路からの化合物の生体利用効率を強化してもよい。最も効果的な剤型は、選択された特定の薬剤の薬物動態、ならびに患者の疾患の重症度に依存するであろう。本開示に従う化合物の鼻内投与、気管内投与もしくは肺投与用のスプレー、ミスト、またはエアロゾルとしての投与が使用されてもよい。したがって本開示はさらに、本明細書に記載の化合物の有効量と、薬剤的に許容可能な担体、添加剤または賦形剤を任意で併用して含む医薬組成物も目的とする。本開示に従う化合物は、即時放出型、中間放出型、または徐放型もしくは制御放出型で投与されてもよい。徐放型または制御放出型は経口投与されることが好ましいが、坐剤および経皮的またはその他の局所型でも投与される。リポソーム型での筋肉内注射を使用して、注射部位での化合物の放出を制御または維持してもよい。 The compounds described herein may be administered in accordance with the present disclosure by oral, parenteral or topical routes in single or divided doses. Administration of the active compound may range from continuous administration (intravenous drip) to several oral doses per day (e.g., Q.I.D.), including oral, topical, parenteral, intramuscular, intravenous, subcutaneous, transdermal (which may include a penetration enhancer), buccal, sublingual, and suppository administration, among other routes of administration. Enteric-coated oral tablets may be used to enhance the bioavailability of the compound from the oral route of administration. The most effective dosage form will depend on the pharmacokinetics of the particular agent selected, as well as the severity of the patient's disease. Administration of the compounds according to the present disclosure as a spray, mist, or aerosol for intranasal, intratracheal, or pulmonary administration may be used. Thus, the present disclosure is further directed to pharmaceutical compositions comprising an effective amount of the compounds described herein, optionally in combination with a pharmaceutically acceptable carrier, additive, or excipient. The compounds according to the present disclosure may be administered in immediate, intermediate, or sustained or controlled release form. Slow or controlled release forms are preferably administered orally, but also via suppositories and transdermal or other topical forms. Intramuscular injections in liposomal form may also be used to control or sustain the release of the compound at the injection site.
本明細書に記載の組成物は、1つ以上の薬学的に許容可能な担体を使用して従来的な方法で製剤化されてもよく、および制御放出製剤で投与されてもよい。これらの医薬組成物中で使用され得る薬学的に許容可能な担体としては、限定されないが、イオン交換剤、アルミナ、ステアリン酸アルミニウム、レシチン、血清タンパク質、例えばヒト血清アルブミン、緩衝物質、例えばリン酸塩、グリシン、ソルビン酸、ソルビン酸カリウム、飽和植物性脂肪酸の部分グリセリド混合物、水、塩または電解質、例えば硫酸プロラミン、リン酸水素二ナトリウム、リン酸水素カリウム、塩化ナトリウム、亜鉛塩、コロイド状シリカ、三ケイ酸マグネシウム、ポリビニルピロリドン、セルロース系物質、ポリエチレングリコール、カルボキシメチルセルロースナトリウム、ポリアクリル酸塩、ワックス、ポリエチレン-ポリオキシプロピレンブロックポリマー、ポリエチレングリコール、および羊毛脂が挙げられる。 The compositions described herein may be formulated in a conventional manner using one or more pharma- ceutically acceptable carriers and may be administered in controlled release formulations. Pharmaceutically acceptable carriers that may be used in these pharmaceutical compositions include, but are not limited to, ion exchangers, alumina, aluminum stearate, lecithin, serum proteins such as human serum albumin, buffer substances such as phosphates, glycine, sorbic acid, potassium sorbate, partial glyceride mixtures of saturated vegetable fatty acids, water, salts or electrolytes such as prolamin sulfate, disodium hydrogen phosphate, potassium hydrogen phosphate, sodium chloride, zinc salts, colloidal silica, magnesium trisilicate, polyvinylpyrrolidone, cellulosic substances, polyethylene glycol, sodium carboxymethylcellulose, polyacrylates, waxes, polyethylene-polyoxypropylene block polymers, polyethylene glycol, and wool fat.
本明細書に記載の組成物は、経口的に、吸入スプレーにより非経口的に、局所的に、直腸内に、鼻内に、口腔内に、膣内に、または移植容器を介して投与されてもよい。本明細書で使用される「非経口」という用語は、皮下、静脈内、筋肉内、関節内、滑膜内、胸骨内、くも膜下腔内、肝臓内、病変内、および頭蓋内の注射または点滴技術を含む。組成物は、経口、腹腔内、または静脈内に投与されることが好ましい。 The compositions described herein may be administered orally, parenterally by inhalation spray, topically, rectally, nasally, buccally, vaginally, or via an implanted vessel. As used herein, the term "parenteral" includes subcutaneous, intravenous, intramuscular, intra-articular, intra-synovial, intrasternal, intrathecal, intrahepatic, intralesional, and intracranial injection or infusion techniques. The compositions are preferably administered orally, intraperitoneally, or intravenously.
本明細書に記載の組成物の滅菌注射形態は、水性懸濁液また油性懸濁液であってもよい。これらの懸濁液は、適切な分散剤または湿潤剤と、懸濁化剤を使用して、当分野で公知の技術より製剤化されてもよい。滅菌注射用調製物は、例えば1,3-ブタンジオール溶液としてなど、非毒性の非経口的に許容可能な希釈剤または溶媒での滅菌注射用溶液または懸濁液であってもよい。採用され得る許容可能なビヒクルおよび溶媒の中では、水、リンゲル溶液と等張塩化ナトリウム溶液がある。加えて滅菌された固定油が溶媒または懸濁媒体として慣例的に使用される。この目的に対し、合成モノグリセリドまたはジグリセリドを含む任意のブランドの固定油を採用してもよい。例えばオリーブオイルまたはヒマシ油、特にそれらのポリオキシエチル化型など、天然の薬学的に許容可能な油があるように、例えばオレイン酸およびそのグリセリド誘導体などの脂肪酸は、注射剤の調製に有用である。これらの油性溶液または懸濁液は、長鎖アルコール希釈剤または分散剤、例えばPh.Helvまたは類似アルコールなどを含んでもよい。 Sterile injectable forms of the compositions described herein may be aqueous or oleaginous suspensions. These suspensions may be formulated by techniques known in the art using suitable dispersing or wetting agents and suspending agents. Sterile injectable preparations may be sterile injectable solutions or suspensions in a non-toxic parenterally acceptable diluent or solvent, for example as a solution in 1,3-butanediol. Among the acceptable vehicles and solvents that may be employed are water, Ringer's solution, and isotonic sodium chloride solution. In addition, sterile fixed oils are conventionally used as solvents or suspending media. For this purpose, any bland fixed oil may be employed, including synthetic mono- or diglycerides. Fatty acids such as oleic acid and its glyceride derivatives are useful in the preparation of injectables, as are natural pharma-ceutically acceptable oils, such as olive oil or castor oil, especially in their polyoxyethylated versions. These oil solutions or suspensions may contain long-chain alcohol diluents or dispersants, such as Ph. Helv or similar alcohols.
本明細書に記載の医薬組成物は、限定されないがカプセル、錠剤、水性懸濁液または水溶液をはじめとする任意の経口的に許容可能な剤形で経口投与されてもよい。経口使用のための錠剤の場合、一般的に使用される担体としてはラクトースおよびコーンスターチが挙げられる。典型的には例えばステアリン酸マグネシウムなどの潤滑剤も添加される。カプセル型での経口投与については、有用な希釈剤としてはラクトースおよび乾燥コーンスターチが挙げられる。経口使用に水性懸濁液が必要な場合、活性成分は乳化剤および懸濁化剤と組み合わされる。望ましい場合、特定の甘味剤、香味剤、または着色剤も添加されてもよい。 The pharmaceutical compositions described herein may be orally administered in any orally acceptable dosage form, including, but not limited to, capsules, tablets, aqueous suspensions or solutions. In the case of tablets for oral use, commonly used carriers include lactose and cornstarch. Lubricants, such as magnesium stearate, are also typically added. For oral administration in capsule form, useful diluents include lactose and dried cornstarch. When aqueous suspensions are required for oral use, the active ingredient is combined with emulsifying and suspending agents. Certain sweetening, flavoring, or coloring agents may also be added, if desired.
あるいは、本明細書に記載の医薬組成物は、直腸投与用の坐剤の形態で投与されてもよい。これらは、室温で固体であるが直腸温度で液体となり、したがって、直腸で溶融して薬物を放出するであろう、適切な非刺激性の賦形剤と混合することによって調製されることができる。かかる物質としては、ココアバター、蜜ろうおよびポリエチレングリコールが挙げられる。 Alternatively, the pharmaceutical compositions described herein may be administered in the form of suppositories for rectal administration. These can be prepared by mixing with a suitable non-irritating excipient that is solid at room temperature but liquid at rectal temperature and will therefore melt in the rectum and release the drug. Such materials include cocoa butter, beeswax and polyethylene glycols.
また本明細書に記載の医薬組成物は、局所的に投与されてもよい。適切な局所用製剤は、これらの各領域または器官のそれぞれに対して容易に調製される。下部腸管への局所適用は、直腸坐剤製剤(上記参照)または適切な浣腸剤で効き目をもたらすことができる。局所的に許容可能な経皮パッチを使用してもよい。 The pharmaceutical compositions described herein may also be administered topically. Suitable topical formulations are readily prepared for each of these areas or organs. Topical application to the lower intestinal tract may be effected in a rectal suppository formulation (see above) or in a suitable enema formulation. Topically acceptable transdermal patches may also be used.
局所適用に関し、医薬組成物は、1つ以上の担体中に懸濁または溶解された活性成分を含有する適切な軟膏で製剤化されてもよい。本開示化合物の局所投与用の担体としては限定されないが鉱油、流動ワセリン、白色ワセリン、プロピレングリコール、ポリオキシエチレン、ポリオキシプロピレン化合物、乳化ろうおよび水が挙げられる。本開示の特定の好ましい態様では、化合物は、患者に外科的に移植されるステント上にコーティングされてもよく、それにより患者内のステントで閉塞が発生する可能性が抑制または低減される。 For topical application, the pharmaceutical composition may be formulated in a suitable ointment containing the active ingredient suspended or dissolved in one or more carriers. Carriers for topical administration of the disclosed compounds include, but are not limited to, mineral oil, liquid petrolatum, white petrolatum, propylene glycol, polyoxyethylene, polyoxypropylene compounds, emulsifying wax, and water. In certain preferred aspects of the present disclosure, the compounds may be coated onto a stent that is surgically implanted in a patient, thereby preventing or reducing the likelihood of occlusion occurring at the stent in the patient.
あるいは医薬組成物は、1つ以上の薬学的に許容可能な担体中に懸濁または溶解された活性成分を含有する適切なローションまたはクリームで製剤化されてもよい。適切な担体としては、限定されないが、鉱油、モノステアリン酸ソルビタン、ポリソルベート60、セチルエステルワックス、セテアリルアルコール、2-オクチルドデカノール、ベンジルアルコールおよび水が挙げられる。 Alternatively, the pharmaceutical compositions may be formulated in a suitable lotion or cream containing the active ingredients suspended or dissolved in one or more pharma- ceutically acceptable carriers. Suitable carriers include, but are not limited to, mineral oil, sorbitan monostearate, polysorbate 60, cetyl esters wax, cetearyl alcohol, 2-octyldodecanol, benzyl alcohol and water.
眼科用途に対し、医薬組成物は、等張性で、pH調整された滅菌生理食塩水の微粉化懸濁液として、または好ましくは等張性でpH調整された滅菌生理食塩水の溶液として、製剤化されてもよく、いずれも例えば塩化ベンジルアルコニウムなどの防腐剤の有無は問わない。あるいは眼科用途に対し、医薬組成物は、例えばワセリンなどの軟膏で製剤化されてもよい。 For ophthalmic applications, the pharmaceutical composition may be formulated as a micronized suspension in isotonic, pH-adjusted, sterile saline, or preferably as a solution in isotonic, pH-adjusted, sterile saline, either with or without a preservative, e.g., benzylalkonium chloride. Alternatively, for ophthalmic applications, the pharmaceutical composition may be formulated in an ointment, e.g., petrolatum.
本明細書に記載の医薬組成物は鼻エアロゾルまたは吸入により投与されてもよい。そのような組成物は、医薬製剤分野に公知の技術に従って調製され、そして生理食塩水の溶液として、ベンジルアルコールまたはその他の適切な防腐剤、生体利用効率を向上させるための吸収プロモーター、フルオロカーボン、および/またはその他の慣用的な可溶化剤または分散剤を採用して調製されてもよい。 The pharmaceutical compositions described herein may be administered by nasal aerosol or inhalation. Such compositions may be prepared according to techniques known in the pharmaceutical formulation art, and may be prepared as solutions in saline, employing benzyl alcohol or other suitable preservatives, absorption promoters to enhance bioavailability, fluorocarbons, and/or other conventional solubilizing or dispersing agents.
単一剤形を生成するために担体材料と合わせられ得る本明細書に記載される医薬組成物中の化合物の量は、治療される宿主および疾患、ならびに特定の投与様式によって異なるであろう。好ましくは、組成物は、単独で、または本開示に従う少なくとも1種の他の化合物と併用して、約0.05ミリグラム~約750ミリグラム以上、より好ましくは約1ミリグラム~約600ミリグラム、さらにより好ましくは約10ミリグラム~約500ミリグラムの活性成分を含んで製剤化されるべきである。 The amount of the compounds in the pharmaceutical compositions described herein that can be combined with the carrier materials to produce a single dosage form will vary depending on the host and disease being treated, as well as the particular mode of administration. Preferably, the compositions should be formulated to contain about 0.05 milligrams to about 750 milligrams or more of active ingredient, either alone or in combination with at least one other compound according to the present disclosure, more preferably about 1 milligram to about 600 milligrams, and even more preferably about 10 milligrams to about 500 milligrams.
任意の特定の患者に対する具体的な投与量および治療レジメンは、使用される具体的な化合物の活性、年齢、体重、一般健康状態、性別、食事、投与時間、排泄速度、薬剤の組み合わせ、および治療を行う医師の判断、および治療を受ける特定の疾患または状態の重症度をはじめとする様々な因子に依存することも理解されたい。 It should also be understood that the specific dosage and treatment regimen for any particular patient will depend on a variety of factors, including the activity of the specific compound used, the age, body weight, general health, sex, diet, time of administration, rate of excretion, drug combination, and the judgment of the treating physician, as well as the severity of the particular disease or condition being treated.
本明細書に記載の方法に従う化合物を使用した治療を必要とする患者または対象は、その薬学的に許容可能な塩、溶媒和物または多形体を含む本開示に従う化合物の有効量を、任意で薬学的に許容可能な担体または希釈剤中で単独で、または本明細書に別途特定される他の公知の治療剤と併用して、患者(対象)に投与することにより治療されることができる。 A patient or subject in need of treatment with a compound according to the methods described herein can be treated by administering to the patient (subject) an effective amount of a compound according to the present disclosure, including a pharma- ceutically acceptable salt, solvate or polymorph thereof, optionally in a pharma- ceutically acceptable carrier or diluent, alone or in combination with other known therapeutic agents as otherwise specified herein.
これらの化合物は、経口、非経口、静脈内、皮内、皮下、または局所などの任意の適切な経路により投与されることができ、例えば液体、クリーム、ゲル、または固体形態で経皮的に、またはエアロゾル形態により投与されることができる。 These compounds can be administered by any suitable route, such as orally, parenterally, intravenously, intradermally, subcutaneously, or topically, for example, transdermally in liquid, cream, gel, or solid form, or in aerosol form.
所望される適応症に対する治療有効量を患者に送達するのに充分であり、治療される患者に重篤な毒性作用をもたらさない量で、薬学的に許容可能な担体または希釈剤中に活性化合物が含有される。本明細書において言及される状態のすべてに対する活性化合物の好ましい用量は、約10ng/kg~300mg/kgの範囲であり、好ましくは一日当たり0.1~100mg/kgの範囲、より一般的にはレシピエント/患者の体重kg当たり、一日当たり0.5~約25mgの範囲である。典型的な局所投与量は、適切な担体中、0.01~5%wt/wtの範囲でありえる。 The active compound is contained in a pharma- ceutically acceptable carrier or diluent in an amount sufficient to deliver to the patient a therapeutically effective amount for the desired indication, but which does not result in serious toxic effects in the patient being treated. Preferred doses of the active compound for all of the conditions mentioned herein range from about 10 ng/kg to 300 mg/kg, preferably in the range of 0.1 to 100 mg/kg per day, and more typically in the range of 0.5 to about 25 mg per kg of recipient/patient body weight per day. Typical topical dosages can range from 0.01 to 5% wt/wt in a suitable carrier.
化合物は、単位剤形当たり1mg未満、1mg~3000mg、好ましくは5~500mgの活性成分を含有する任意の適切な単位剤形で簡便に投与されるが、これに限定されない。約25~250mgの経口投与量が多くの場合、便利である。 The compounds are conveniently administered in any suitable unit dosage form, including, but not limited to, less than 1 mg, 1 mg to 3000 mg, and preferably 5 to 500 mg of active ingredient per unit dosage form. An oral dosage of about 25 to 250 mg is often convenient.
活性成分は、約0.00001~30mM、好ましくは約0.1~30μMの活性化合物のピーク血漿濃度を実現するよう投与されることが好ましい。これは、例えば、活性成分の溶液または製剤を、任意で生理食塩水溶液中、または水性媒体中で静脈内注射を行うことにより、または活性成分のボーラス投与により実現し得る。経口投与は、活性薬剤の有効血漿濃度を生じさせるためにも適している。 The active ingredient is preferably administered to achieve a peak plasma concentration of the active compound of about 0.00001-30 mM, preferably about 0.1-30 μM. This may be achieved, for example, by intravenous injection of a solution or formulation of the active ingredient, optionally in saline solution or in an aqueous medium, or by bolus administration of the active ingredient. Oral administration is also suitable for producing effective plasma concentrations of the active agent.
薬物組成物中の活性化合物の濃度は、薬剤の吸収、分布、不活性化、および排出速度、ならびに当分野の当業者に公知の他の因子に依存するであろう。投薬量の値は、軽減される状態の重大度によっても変化するであろうことに留意されたい。さらには、任意の特定の対象に対して、個人の要求、および当該組成物の投与を行い、または投与を管理する人物の専門的な判断に従い特定の投薬レジメンが経時的に調整されるべきであること、また本明細書に記載される濃度範囲は単なる例示であり、特許請求される組成物の範囲または実施を限定することは意図されていないことを理解されたい。活性成分は、一度に投与されてもよく、または多数のより少ない用量に分割されて様々な時間間隔で投与されてもよい。 The concentration of the active compound in the drug composition will depend on the absorption, distribution, inactivation, and excretion rates of the drug, as well as other factors known to those skilled in the art. It is noted that dosage values will also vary with the severity of the condition to be alleviated. It is further understood that for any particular subject, specific dosage regimens should be adjusted over time according to the individual needs and the professional judgment of the person administering or supervising the administration of the composition, and that the concentration ranges set forth herein are merely exemplary and are not intended to limit the scope or practice of the claimed compositions. The active ingredient may be administered at once, or may be divided into multiple smaller doses to be administered at various time intervals.
経口組成物は一般に、不活性希釈剤または食用担体を含むであろう。ゼラチンカプセルで封入されてもよく、または錠剤に圧縮されてもよい。経口治療投与の目的で、活性化合物またはそのプロドラッグ誘導体を賦形剤と組み合わせ、錠剤、トローチ、またはカプセルの形態で使用することができる。薬学的に適合性のある結合剤および/またはアジュバント物質を組成物の一部として含有させることができる。 Oral compositions will generally include an inert diluent or an edible carrier. They may be enclosed in gelatin capsules or compressed into tablets. For the purpose of oral therapeutic administration, the active compound or its prodrug derivative can be combined with excipients and used in the form of tablets, troches, or capsules. Pharmaceutically compatible binding agents, and/or adjuvant materials can be included as part of the composition.
錠剤、丸剤、カプセル、トローチなどは、以下の成分:微結晶性セルロース、トラガカントガム、もしくはゼラチンなどの結合剤;デンプンもしくはラクトースなどの賦形剤;アルギン酸、プリモゲル、もしくはトウモロコシデンプンなどの分散剤;ステアリン酸マグネシウムもしくはステロテスなどの潤滑剤;コロイド状二酸化ケイ素などの流動剤;スクロースもしくはサッカリンなどの甘味剤;またはペパーミント、サリチル酸メチル、もしくはオレンジ香味料などの香味剤、あるいはそれらと同様の性質の化合物のうちのいずれかを含有することができる。投薬単位剤型がカプセルである場合、上述のタイプの物質に加えて例えば脂肪酸などの液状担体を含むことができる。さらに、投薬単位剤型は、例えば、糖コーティング、セラック、または腸溶剤などの物理的な投薬単位の物理的形状を改変する様々な他の物質を含有することができる。 Tablets, pills, capsules, troches, etc. may contain any of the following ingredients: binders such as microcrystalline cellulose, tragacanth, or gelatin; excipients such as starch or lactose; dispersants such as alginic acid, primogel, or corn starch; lubricants such as magnesium stearate or sterotes; flow agents such as colloidal silicon dioxide; sweeteners such as sucrose or saccharin; or flavorings such as peppermint, methyl salicylate, or orange flavoring, or compounds of a similar nature. When the dosage unit is a capsule, it may contain, in addition to the above types of materials, a liquid carrier such as, for example, a fatty acid. In addition, the dosage unit may contain various other materials that modify the physical form of the dosage unit, such as, for example, sugar coatings, shellac, or enteric agents.
活性化合物またはその薬学的に許容可能な塩は、エリキシル、懸濁液、シロップ、ウェハース、チューインガムなどの構成要素として投与されることができる。シロップは、活性化合物に加えて、甘味剤としてスクロース、あるいは特定の防腐剤、染料および着色剤、ならびに香味料を含んでもよい。 The active compound or a pharma- ceutically acceptable salt thereof can be administered as a component of an elixir, suspension, syrup, wafer, chewing gum or the like. A syrup may contain, in addition to the active compound, sucrose as a sweetening agent and/or certain preservatives, dyes and colorings, and flavors.
活性化合物またはその薬学的に許容される塩は、所望の作用を損なわない他の活性物質、または所望の作用を補完する物質、例えば、とりわけ、ペムブロリズマブを含む抗がん剤と混合することもできる。本開示の特定の好ましい態様において、本開示に従う1つ以上の化合物は、本明細書に別段に記載される例えば抗がん剤、または抗生物質を含む創傷治癒剤などの別の生物活性薬剤とともに共投与される。 The active compounds or pharma- ceutically acceptable salts thereof may also be mixed with other active substances that do not impair the desired action or that complement the desired action, such as anti-cancer drugs, including pembrolizumab, among others. In certain preferred aspects of the present disclosure, one or more compounds according to the present disclosure are co-administered with another bioactive agent, such as an anti-cancer drug, or a wound healing agent, including an antibiotic, as described elsewhere herein.
非経口、皮内、皮下、または局所適用に使用される溶液または懸濁液には、以下の構成要素が含まれてもよい:例えば注射用水、生理食塩水溶液、固定油、ポリエチレングリコール、グリセリン、プロピレングリコール、またはその他の合成溶媒などの滅菌希釈剤;例えばベンジルアルコールまたはメチルパラベンなどの抗菌剤;例えばアスコルビン酸または重亜硫酸ナトリウムなどの抗酸化剤;例えばエチレンジアミン四酢酸などのキレート剤;例えば酢酸塩、クエン酸塩、またはリン酸塩などの緩衝剤、および例えば塩化ナトリウムまたはデキストロースなどの張力調整用の剤。非経口調製物は、ガラスまたはプラスチックで作製されたアンプル、使い捨てシリンジ、または複数投与用バイアルに封入されてもよい。 Solutions or suspensions used for parenteral, intradermal, subcutaneous, or topical application may include the following components: a sterile diluent such as water for injection, saline solution, fixed oils, polyethylene glycols, glycerin, propylene glycol, or other synthetic solvents; antibacterial agents such as benzyl alcohol or methylparabens; antioxidants such as ascorbic acid or sodium bisulfite; chelating agents such as ethylenediaminetetraacetic acid; buffers such as acetates, citrates, or phosphates, and agents for adjusting tonicity such as sodium chloride or dextrose. Parenteral preparations may be enclosed in ampoules, disposable syringes, or multiple dose vials made of glass or plastic.
静脈内投与された場合、好ましい担体は生理食塩水またはリン酸緩衝生理食塩水(PBS)である。 When administered intravenously, the preferred carrier is saline or phosphate buffered saline (PBS).
1つの実施形態では、活性化合物は、例えばインプラントおよびマイクロカプセル化送達システムをはじめとする放出制御製剤など、身体からの急速な排出から化合物を保護する担体で調製される。例えばエチレンビニル酢酸塩、ポリ無水物、ポリグリコール酸、コラーゲン、ポリオルトエステル、およびポリ乳酸など、生分解性で生体適合性のポリマーを使用することができる。こうした製剤の調製方法は、当分野の当業者には明らかであろう。 In one embodiment, the active compounds are prepared with carriers that will protect the compound against rapid elimination from the body, such as a controlled release formulation, including implants and microencapsulated delivery systems. Biodegradable, biocompatible polymers can be used, such as ethylene vinyl acetate, polyanhydrides, polyglycolic acid, collagen, polyorthoesters, and polylactic acid. Methods for preparing such formulations will be apparent to those skilled in the art.
リポソーム懸濁液も薬学的に許容可能な担体であり得る。これらは、当業者に既知である方法、例えば、米国特許第4,522,811号(参照によりその全体が本明細書に組み込まれる)に記載される方法に従って調製され得る。例えば、リポソーム製剤は、適切な脂質(例えばステアロイルホスファチジルエタノールアミン、ステアロイルホスファチジルコリン、アラカドイル(arachadoyl)ホスファチジルコリン、およびコレステロールなど)を無機溶媒中で溶解させ、次いで蒸発させて、容器表面上に乾燥した脂質の薄膜を残すことにより調製されてもよい。次いで活性化合物の水溶液を容器に入れる。次いで容器を手で回転させ、容器の側面から脂質材料を剥がし、脂質塊を分散させ、リポソーム懸濁液を形成させる。 Liposomal suspensions may also be pharma- ceutically acceptable carriers. These may be prepared according to methods known to those skilled in the art, such as those described in U.S. Pat. No. 4,522,811, which is incorporated herein by reference in its entirety. For example, liposomal formulations may be prepared by dissolving appropriate lipids (e.g., stearoyl phosphatidylethanolamine, stearoyl phosphatidylcholine, arachadoyl phosphatidylcholine, and cholesterol, etc.) in an inorganic solvent, which is then evaporated to leave a thin film of dried lipid on the container surface. An aqueous solution of the active compound is then placed in the container. The container is then rotated by hand to remove lipid material from the sides of the container, dispersing the lipid mass and forming the liposomal suspension.
治療方法
追加の態様において本明細書は、本明細書に記載の化合物またはその塩型の有効量、および薬学的に許容可能な担体を含む治療用組成物を提供する。治療用組成物は、患者または対象、例えばヒトなどの動物においてタンパク質の分解を調節し、当該分解されたタンパク質を介して調節される病状もしくは状態の治療または改善に使用することができる。
In an additional aspect, the present specification provides a therapeutic composition comprising an effective amount of a compound described herein or a salt form thereof, and a pharma- ceutically acceptable carrier, which can be used to modulate protein degradation in a patient or subject, e.g., an animal, such as a human, and to treat or ameliorate a disease state or condition that is modulated via the degraded protein.
本明細書において使用される場合、「治療する」、「治療すること」、および「治療」という用語は、患者に利益をもたらす何らかの作用を指し、本化合物が結合するタンパク質を介して調節される任意の病態または状態の治療を含む、その利益のために本化合物が投与され得る。本開示に従う化合物を使用して治療され得る、非小細胞肺がんを含む肺がんなどのがんを含む疾患状態または状態は、本明細書の上記に記載されている。 As used herein, the terms "treat," "treating," and "treatment" refer to any action that benefits a patient, for which the compounds may be administered, including the treatment of any disease state or condition that is modulated via the protein to which the compounds bind. Disease states or conditions, including cancers such as lung cancer, including non-small cell lung cancer, that may be treated using compounds according to the present disclosure are described herein above.
本明細書は、疾患、例えば、がんの治療または改善を目的として対象タンパク質の分解を生じさせるための、本明細書に記載の治療用組成物を提供する。特定の追加的実施形態において、疾患は多発性骨髄腫である。したがって、別の態様において本明細書は、細胞中の標的タンパク質をユビキチン化/分解する方法を提供する。ある特定の実施形態では、本方法は、本明細書に記載される二官能性化合物を投与することを含み、化合物は、例えば、本明細書に別途記載されるように、好ましくはリンカー部分を介して結合された、ULMおよびPTMを含み、ULMはPTMに結合し、かつULMは、ユビキチン経路タンパク質(例えば、VHL E3ユビキチンリガーゼなどのユビキチンリガーゼ)を認識し、PTMは、標的タンパク質がユビキチンリガーゼに近接して配置されると標的タンパク質の分解が生じ、ひいては、標的タンパク質の効果の分解/阻害およびタンパク質レベルの制御をもたらすように、標的タンパク質を認識する。本開示によりもたらされるタンパク質レベルの制御は病態または状態の治療を提供するものであり、これは例えば患者細胞などの細胞中のそのタンパク質のレベルを低下させることによって標的タンパク質を介して調節される。特定の実施形態では、本方法は、本明細書に記載の化合物の有効量を、任意で薬学的に許容可能な賦形剤、担体、アジュバント、別の生物活性剤、またはそれらの組み合わせを含み、投与することを含む。 The present disclosure provides therapeutic compositions as described herein for causing degradation of a target protein for the treatment or amelioration of a disease, e.g., cancer. In certain additional embodiments, the disease is multiple myeloma. Thus, in another aspect, the present disclosure provides a method for ubiquitinating/degrading a target protein in a cell. In certain embodiments, the method includes administering a bifunctional compound as described herein, the compound including a ULM and a PTM, preferably linked via a linker moiety, e.g., as described elsewhere herein, where the ULM binds to the PTM, and the ULM recognizes a ubiquitin pathway protein (e.g., a ubiquitin ligase, such as VHL E3 ubiquitin ligase), and the PTM recognizes the target protein such that when the target protein is placed in proximity to the ubiquitin ligase, degradation of the target protein occurs, thus resulting in degradation/inhibition of the effect of the target protein and control of the protein level. The control of protein level provided by the present disclosure provides treatment of a disease state or condition that is regulated via the target protein by reducing the level of that protein in a cell, e.g., a patient cell. In certain embodiments, the method includes administering an effective amount of a compound described herein, optionally including a pharma- ceutically acceptable excipient, carrier, adjuvant, another bioactive agent, or a combination thereof.
追加的実施態様において本明細書は、例えばヒトなどの動物などの対象または患者において、疾患、障害もしくはその症状を治療または改善する方法を提供するものであり、当該方法は、本明細書に記載の化合物またはその塩型の治療有効量などの有効量と、薬学的に許容可能な賦形剤、担体、アジュバント、別の生物活性剤、またはそれらの組み合わせを含む組成物をその必要のある対象に投与することを含み、この場合において当該組成物は、対象において疾患もしくは障害またはその症状の治療または改善に有効である。 In additional embodiments, the present specification provides a method of treating or ameliorating a disease, disorder, or a symptom thereof in a subject or patient, e.g., an animal, e.g., a human, comprising administering to a subject in need thereof a composition comprising an effective amount, e.g., a therapeutically effective amount, of a compound described herein, or a salt form thereof, and a pharma- ceutically acceptable excipient, carrier, adjuvant, another bioactive agent, or a combination thereof, wherein the composition is effective to treat or ameliorate a disease or disorder, or a symptom thereof, in the subject.
別の態様において本明細書は、本開示による化合物を使用して生物的システムにおいて対象タンパク質の分解効果を特定する方法を提供する。 In another aspect, the present specification provides a method for identifying the degradative effect of a protein of interest in a biological system using a compound according to the present disclosure.
別の実施形態において、本開示は、タンパク質を介して調節される病態または状態に関して必要のあるヒト患者を治療する方法を目的としており、ここで当該タンパク質の分解は当該患者において治療効果を生じさせるものであり、当該方法は、本開示化合物の有効量を、任意で別の生物活性剤と併用して必要のある患者に投与することを含む。病態または状態は、微生物体、または例えばウイルス、細菌、真菌、原虫もしくはその他の微生物などの他の外来性主体により引き起こされる疾患であってもよく、または病態および/もしくは状態が生じるタンパク質の過剰発現により引き起こされる病態であってもよい。 In another embodiment, the disclosure is directed to a method of treating a human patient in need for a disease state or condition regulated through a protein, where degradation of the protein produces a therapeutic effect in the patient, comprising administering to a patient in need thereof an effective amount of a compound of the disclosure, optionally in combination with another bioactive agent. The disease state or condition may be a disease caused by a microbial organism or other foreign agent, such as, for example, a virus, a bacterium, a fungus, a protozoan, or other microorganism, or may be a disease state caused by overexpression of a protein resulting in the disease state and/or condition.
「病態または状態」という用語は、タンパク質の調節異常(すなわち、患者で発現されているタンパク質の量が増加している)が発生し、患者中の1つ以上のタンパク質の分解が、それを必要とする患者に有益な治療または症状の軽減をもたらす任意の病態または状態を記載するために使用される。特定の例では、病態または状態は治療され得る。 The term "disease state or condition" is used to describe any disease state or condition in which dysregulation of a protein occurs (i.e., the amount of a protein expressed in a patient is increased) and degradation of one or more proteins in the patient provides beneficial treatment or relief of symptoms to a patient in need thereof. In certain instances, the disease state or condition may be treated.
本開示に従う化合物を使用して治療し得る疾患状態または状態には、例えば、喘息、自己免疫疾患、例えば、多発性硬化症、様々ながん、繊毛関連疾患、口蓋裂、糖尿病、心臓病、高血圧、炎症性腸疾患、精神遅滞、気分障害、肥満、屈折異常、不妊症、アンジェルマン症候群、カナバン病、セリアック病、シャルコー・マリー・トゥース病、嚢胞性線維症、デュシェンヌ筋ジストロフィー、血色素症、血友病、クラインフェルター症候群、神経線維腫症、フェニルケトン尿症、多発性嚢胞腎疾患、(PKD1)または4(PKD2)プラダー・ウィリー症候群、鎌状赤血球症、テイ・サックス病、ターナー症候群が挙げられる。 Disease states or conditions that may be treated using compounds according to the present disclosure include, for example, asthma, autoimmune diseases, such as multiple sclerosis, various cancers, cilia-related diseases, cleft palate, diabetes, heart disease, hypertension, inflammatory bowel disease, mental retardation, mood disorders, obesity, refractive errors, infertility, Angelman syndrome, Canavan disease, celiac disease, Charcot-Marie-Tooth disease, cystic fibrosis, Duchenne muscular dystrophy, hemochromatosis, hemophilia, Klinefelter syndrome, neurofibromatosis, phenylketonuria, polycystic kidney disease, (PKD1) or 4 (PKD2) Prader-Willi syndrome, sickle cell disease, Tay-Sachs disease, Turner syndrome.
「新生物」または「がん」という用語は本明細書全体を通して、がん性または悪性の新生物、すなわち、多くの場合正常よりもずっと早く細胞が増殖することにより成長し、開始されたその新たな成長を止める刺激の後にも成長し続ける異常な組織の形成および成長をもたらす病的プロセスを指すために使用される。悪性新生物は、構造的組織化の部分的または完全な欠落、および正常組織との機能的協調の欠落を示し、また大部分が周辺組織へ侵入し、いくつかの部位へと転移し、除去が試みられた後にも再発し、適切に治療されない限り患者が死亡する確率が高い。本明細書において使用される場合、新生物という用語は、すべてのがん性病態を記載するために使用され、悪性造血細胞、腹水および固形腫瘍と関連する病的プロセスを包含する。本化合物単独で、または少なくとも一種の追加的抗がん剤と併用して本化合物により治療され得るがんの例としては、扁平上皮細胞がん、基底細胞がん、腺がん、肝細胞がんおよび腎細胞がん、膀胱がん、腸がん、乳がん、子宮頸がん、大腸がん、食道がん、頭部がん、腎がん、肝がん、肺がん、頸部がん、卵巣がん、膵がん、前立腺がんおよび胃がん;白血病;良性および悪性のリンパ腫、特にバーキットリンパ腫および非ホジキンリンパ腫;良性および悪性の黒色腫;骨髄増殖性疾患;ユーイング肉腫、血管肉腫、カポジ肉腫、脂肪肉腫、筋肉腫、末梢神経腫、滑膜肉腫、グリオーマ、星膠細胞腫、乏突起膠腫、上衣腫、グリア芽腫、神経芽細胞腫、神経節細胞腫、神経節膠腫、髄芽細胞腫、松果体細胞腫瘍、髄膜腫、髄膜肉腫、神経線維腫、および神経鞘腫を含む肉腫;大腸がん、乳がん、前立腺がん、子宮頸がん、子宮がん、肺がん、卵巣がん、精巣がん、甲状腺がん、星状細胞腫、食道がん、膵がん、胃がん、肝がん、大腸がん、黒色腫;がん肉腫、ホジキン病、ウィルムス腫瘍および奇形がんが挙げられる。本開示に従う化合物を使用して治療され得るさらなるがんとしては、例えば、T細胞系急性リンパ芽球性白血病(T-ALL)、T細胞系リンパ芽球性リンパ腫(T-LL)、末梢T細胞リンパ腫、成人T細胞リンパ腫、前駆B細胞ALL、前駆B細胞リンパ腫、大細胞型B細胞リンパ腫、バーキットリンパ腫、B細胞性ALL、フィラデルフィア染色体陽性ALL、およびフィラデルフィア染色体陽性CMLが挙げられる。 The term "neoplasm" or "cancer" is used throughout this specification to refer to a cancerous or malignant neoplasm, i.e., a pathological process that results in the formation and growth of abnormal tissue that grows by cell proliferation, often much faster than normal, and continues to grow after stimuli that would have stopped the new growth that was initiated. Malignant neoplasms exhibit partial or complete lack of structural organization and functional coordination with normal tissue, and most invade surrounding tissues, metastasize to several sites, recur after attempted removal, and have a high probability of causing death to the patient unless appropriately treated. As used herein, the term neoplasm is used to describe all cancerous conditions and includes pathological processes associated with malignant hematopoietic cells, ascites, and solid tumors. Examples of cancers that may be treated by the present compounds alone or in combination with at least one additional anti-cancer agent include squamous cell carcinoma, basal cell carcinoma, adenocarcinoma, hepatocellular and renal cell carcinoma, bladder cancer, intestinal cancer, breast cancer, cervical cancer, colon cancer, esophageal cancer, head cancer, kidney cancer, liver cancer, lung cancer, neck cancer, ovarian cancer, pancreatic cancer, prostate cancer and stomach cancer; leukemia; benign and malignant lymphomas, particularly Burkitt's lymphoma and non-Hodgkin's lymphoma; benign and malignant melanoma; myeloproliferative disorders; Ewing's sarcoma, hematopoietic cancer, and hematopoietic cancer. Sarcomas, including ductal sarcoma, Kaposi's sarcoma, liposarcoma, myosarcoma, peripheral neuroma, synovial sarcoma, glioma, astrocytoma, oligodendroglioma, ependymoma, glioblastoma, neuroblastoma, ganglioneuroma, ganglioglioma, medulloblastoma, pineal cell tumor, meningioma, meningeal sarcoma, neurofibroma, and schwannoma; colorectal cancer, breast cancer, prostate cancer, cervical cancer, uterine cancer, lung cancer, ovarian cancer, testicular cancer, thyroid cancer, astrocytoma, esophageal cancer, pancreatic cancer, gastric cancer, liver cancer, colorectal cancer, melanoma; carcinosarcoma, Hodgkin's disease, Wilms' tumor, and teratocarcinoma. Additional cancers that may be treated using compounds according to the present disclosure include, for example, T-cell acute lymphoblastic leukemia (T-ALL), T-cell lymphoblastic lymphoma (T-LL), peripheral T-cell lymphoma, adult T-cell lymphoma, precursor B-cell ALL, precursor B-cell lymphoma, large B-cell lymphoma, Burkitt's lymphoma, B-cell ALL, Philadelphia chromosome positive ALL, and Philadelphia chromosome positive CML.
「生物活性剤」という用語は、本開示に従う化合物以外の剤を記載するために使用され、本化合物が使用された目的の治療、阻害、および/または防除/予防の効果を補助するために、生物活性を有する剤として本化合物と併用して使用される。本明細書の使用に好ましい生物活性薬剤は、本化合物が使用され、または投与される活性と類似した薬理活性を有する剤が挙げられ、例えば抗がん剤、抗ウイルス剤、特に抗HIV剤および抗HCV剤、抗微生物剤、抗真菌剤などが挙げられる。 The term "bioactive agent" is used to describe an agent other than the compounds according to the present disclosure, which is used in combination with the present compounds as an agent having biological activity to aid in the therapeutic, inhibitory, and/or preventative/prophylactic effects for which the present compounds are used. Preferred bioactive agents for use herein include agents having pharmacological activity similar to that for which the present compounds are used or administered, such as anti-cancer agents, anti-viral agents, particularly anti-HIV and anti-HCV agents, anti-microbial agents, anti-fungal agents, and the like.
「追加的抗がん剤」という用語は、がんを治療するために本開示に従う化合物と併用され得る抗がん剤を記述するために使用される。これらの薬剤としては、例えば、エベロリムス、トラベクテジン、アブラキサン、TLK 286、AV-299、DN-101、パゾパニブ、GSK690693、RTA 744、ON 0910.Na、AZD 6244(ARRY-142886)、AMN-107、TKI-258、GSK461364、AZD 1152、エンザスタウリン、バンデタニブ、ARQ-197、MK-0457、MLN8054、PHA-739358、R-763、AT-9263、FLT-3阻害剤、VEGFR阻害剤、EGFR TK阻害剤、オーロラキナーゼ阻害剤、PIK-1調節剤、Bcl-2阻害剤、HDAC阻害剤、c-MET阻害剤、PARP阻害剤、Cdk阻害剤、EGFR TK阻害剤、IGFR-TK阻害剤、抗HGF抗体、PI3キナーゼ阻害剤、AKT阻害剤、mTORC1/2阻害剤、JAK/STAT阻害剤、チェックポイント-1または2阻害剤、接着斑キナーゼ阻害剤、Mapキナーゼキナーゼ(mek)阻害剤、VEGFトラップ抗体、ペメトレキセド、エルロチニブ、ダサチニブ、ニロチニブ、デカタニブ、パニツムマブ、アムルビシン、オレゴボマブ、Lep-etu、ノラトレキシド、azd2171、バタブリン、オファツムマブ、ザノリムマブ、エドテカリン、テトランドリン、ルビテカン、テスミリフェン、オブリメルセン、チシリムマブ、イピリムマブ、ゴシポール、Bio 111、131-I-TM-601、ALT-110、BIO 140、CC 8490、シレンジタイド、ジャイマテカン、IL13-PE38QQR、INO 1001、IPdR1 KRX-0402、ルカントン、LY317615、ノイラジアブ、ビテスパン、Rta 744、Sdx 102、タランパネル、アトラセンタン、Xr 311、ロミデプシン、ADS-100380、スニチニブ、5-フルオロウラシル、ボリノスタット、エトポシド、ゲムシタビン、ドキソルビシン、リポソーマルドキソルビシン、5’-デオキシ-5-フルオロウリジン、ビンクリンスチン、テモゾロミド、ZK-304709、セリシクリブ;PD0325901、AZD-6244、カペシタビン、L-グルタミン酸、N-[4-[2-(2-アミノ-4,7-ジヒドロ-4-オキソ-1H-ピロロ[2,3-d]ピリミジン-5-イル)エチル]ベンゾイル]-、ジナトリウム塩、七水和物、カンプトテシン、PEG標識化イリノテカン、タモキシフェン、トレミフェンクエン酸塩、アナストロゾール、エキセメスタン、レトロゾール、DES(ジエチルスチルベストロール)、エストラジオール、エストロゲン、結合型エストロゲン、ベバシズマブ、IMC-1C11、CHIR-258);3-[5-(メチルスルホニルピペラジンメチル)-インドイル-キノロン、バタラニブ、AG-013736、AVE-0005、酢酸ゴセレリン、酢酸ロイプロリド、トリプトレリンパモ酸塩、酢酸メドロキシプロゲステロン、カプロン酸ヒドロキシプロゲステロン、酢酸メゲストロール、ラロキシフェン、ビカルタミド、フルタミド、ニルタミド、酢酸メゲストロール、CP-724714;TAK-165、HKI-272、エルロチニブ、ラパタニブ、カネルチニブ、ABX-EGF抗体、アービタックス、EKB-569、PKI-166、GW-572016、ロナファルニブ、BMS-214662、チピファルニブ;アミフォスチン、NVP-LAQ824、スベロイルアナリドヒドロキサム酸、バルプロ酸、トリコスタチンA、FK-228、SU11248、ソラフェニブ、KRN951、アミノグルテチミド、アムサクリン、アナグレリド、L-アスパラギナーゼ、カルメット-ゲラン桿菌(BCG)ワクチン、アドリアマイシン、ブレオマイシン、ブセレリン、ブスルファン、カルボプラチン、カルムスチン、クロラムブシル、シスプラチン、クラドリビン、クロドロネート、シプロテロン、シタラビン、ダカルバジン、ダクチノマイシン、ダウノルビシン、ジエチルスチルベストロール、エピルビシン、フルダラビン、フルドロコルチゾン、フルオキシメステロン、フルタミド、グリベック、ゲムシタビン、ヒドロキシ尿素、イダルビシン、イホスファミド、イマチニブ、ロイプロリド、レバミゾール、ロムスチン、メクロレタミン、メルファラン、6-メルカプトプリン、メスナ、メトトレキサート、ミトマイシン、ミトタン、ミトキサントロン、ニルタミド、オクトレオチド、オキサリプラチン、パミドロネート、ペントスタチン、プリカマイシン、ポルフィマー、プロカルバジン、ラルチトレキセド、リツキシマブ、ストレプトゾシン、テニポシド、テストステロン、サリドマイド、チオグアニン、チオテパ、トレチノイン、ビンデシン、13-シスレチノイン酸、フェニルアラニンマスタード、ウラシルマスタード、エストラムスチン、アルトレタミン、フロクスウリジン、5-デオオキシウリジン、シトシンアラビノシド、6-メルカプトプリン、デオキシコホルマイシン、カルシトリオール、バルルビシン、ミトラマイシン、ビンブラスチン、ビノレルビン、トポテカン、ラゾキシン、マリマスタット、COL-3、ネオバスタット、BMS-275291、スクアラミン、エンドスタチン、SU5416、SU6668、EMD121974、インターロイキン-12、IM862、アンギオスタチン、ビタキシン、ドロロキシフェン、イドキシフェン、スピロノラクトン、フィナステリド、シミチジン、トラスツズマブ、デニロイキンジフチトクス、ゲフィチニブ、ボルテゾミブ、パクリタキセル、クレモフォアフリーパクリタキセル、ドセタキセル、エピチロンB、BMS-247550、BMS-310705、ドロロキシフェン、4-ヒドロキシタモキシフェン、ピペンドキシフェン、ERA-923、アルゾキシフェン、フルベストラント、アコルビフェン、ラソフォキシフェン、イドキシフェン、TSE-424、HMR-3339、ZK186619、トポテカン、PTK787/ZK 222584、VX-745、PD 184352、ラパマイシン、40-O-(2-ヒドロキシエチル)-ラパマイシン、テムシロリムス、AP-23573、RAD001、ABT-578、BC-210、LY294002、LY292223、LY292696、LY293684、LY293646、ウォルトマンニン、ZM336372、L-779,450、PEG-フィルグラスチム、ダルベポエチン、エリスロポエチン、顆粒球コロニー刺激因子、ゾレンドロネート、プレドニゾロン、セツキシマブ、顆粒球単球コロニー刺激因子、ヒストレリン、ペグ化インターフェロンアルファ-2a、インターフェロンアルファ-2a、ペグ化インターフェロンアルファ-2b、インターフェロンアルファ-2b、アザシチジン、PEG-L-アスパラギナーゼ、レナリドミド、ゲムツズマブ、ヒドロコルチゾン、インターロイキン-11、デクスラゾキサン、アレムツズマブ、全トランス型レチノイン酸、ケトコナゾール、インターロイキン-2、メゲストロール、免疫グロブリン、窒素マスタード、メチルプレドニゾロン、イブリツモマブチウキセタン、アンドロゲン、デシタビン、ヘキサメチルメラミン、ベキサロテン、トシツモマブ、三酸化ひ素、コルチゾン、エチドロネート、ミトタン、シクロスポリン、リポソームダウノルビシン、エドウィナ-アスパラギナーゼ、ストロンチウム89、カソピタント、ネツピタント、NK-1受容体アンタゴニスト、パロノセトロン、アプレピタント、ジフェンヒドラミン、ヒドロキシジン、メトクロプラミド、ロラゼパム、アルプラゾラム、ハロペリドール、ドロペリドール、ドロナビノール、デキサメタゾン、メチルプレドニゾロン、プロクロルペラジン、グラニセトロン、オンダンセトロン、ドラセトロン、トロピセトロン、ペグフィルグラスチム、エリスロポエチン、エポエチンアルファ、ダルベポエチンアルファ、およびそれらの混合物が挙げられる。 The term "additional anti-cancer agent" is used to describe anti-cancer agents that may be combined with compounds according to the present disclosure to treat cancer. These agents include, for example, everolimus, trabectedin, abraxane, TLK 286, AV-299, DN-101, pazopanib, GSK690693, RTA 744, ON 0910. Na, AZD 6244 (ARRY-142886), AMN-107, TKI-258, GSK461364, AZD 1152, enzastaurin, vandetanib, ARQ-197, MK-0457, MLN8054, PHA-739358, R-763, AT-9263, FLT-3 inhibitors, VEGFR inhibitors, EGFR TK inhibitors, Aurora kinase inhibitors, PIK-1 regulators, Bcl-2 inhibitors, HDAC inhibitors, c-MET inhibitors, PARP inhibitors, Cdk inhibitors, EGFR TK inhibitors, IGFR-TK inhibitors, anti-HGF antibodies, PI3 kinase inhibitors, AKT inhibitors, mTORC1/2 inhibitors, JAK/STAT inhibitors, checkpoint-1 or 2 inhibitors, focal adhesion kinase inhibitors, Map kinase kinase (mek) inhibitors, VEGF trap antibodies, pemetrexed, erlotinib, dasatinib, nilotinib, decatanib, panitumumab, amrubicin, oregovomab, Lep-etu, nolatrexed, azd2171, vatabulin, ofatumumab, zanolimumab, edotecarin, tetrandrine, rubitecan, tesmilifene, oblimersen, ticilimumab, ipilimumab, gossypol, Bio 111, 131-I-TM-601, ALT-110, BIO 140, CC 8490, cilengitide, gymatecan, IL13-PE38QQR, INO 1001, IPdR1 KRX-0402, lucanton, LY317615, neuradiab, vitespam, Rta 744, Sdx 102, talampanel, atrasentan, Xr 311, romidepsin, ADS-100380, sunitinib, 5-fluorouracil, vorinostat, etoposide, gemcitabine, doxorubicin, liposomal doxorubicin, 5'-deoxy-5-fluorouridine, vincristine, temozolomide, ZK-304709, seliciclib; PD0325901, AZD-6244, capecitabine, L-glutamic acid, N-[4-[2-(2-amino-4,7-dihydro-4-oxo-1H-pyrrolo[2,3-d]pyrimidine-5 -yl)ethyl]benzoyl]-, disodium salt, heptahydrate, camptothecin, PEG-labeled irinotecan, tamoxifen, toremifene citrate, anastrozole, exemestane, letrozole, DES (diethylstilbestrol), estradiol, estrogen, conjugated estrogens, bevacizumab, IMC-1C11, CHIR-258); 3-[5-(methylsulfonylpiperazinemethyl)-indolyl-quinolone, vatalanib, AG-013736, AVE -0005, goserelin acetate, leuprolide acetate, triptorelin pamoate, medroxyprogesterone acetate, hydroxyprogesterone caproate, megestrol acetate, raloxifene, bicalutamide, flutamide, nilutamide, megestrol acetate, CP-724714; TAK-165, HKI-272, erlotinib, lapatanib, canertinib, ABX-EGF antibody, Erbitux, EKB-569, PKI-166, GW-572016, lonafarnib, BMS-21 4662, tipifarnib; amifostine, NVP-LAQ824, suberoylanalide hydroxamic acid, valproic acid, trichostatin A, FK-228, SU11248, sorafenib, KRN951, aminoglutethimide, amsacrine, anagrelide, L-asparaginase, Bacillus Calmette-Guerin (BCG) vaccine, adriamycin, bleomycin, buserelin, busulfan, carboplatin, carmustine, chlorambucil, cisplatin, cladribine, clodrofluidine acetaminophen, cyproterone, cytarabine, dacarbazine, dactinomycin, daunorubicin, diethylstilbestrol, epirubicin, fludarabine, fludrocortisone, fluoxymesterone, flutamide, Gleevec, gemcitabine, hydroxyurea, idarubicin, ifosfamide, imatinib, leuprolide, levamisole, lomustine, mechlorethamine, melphalan, 6-mercaptopurine, mesna, methotrexate, mitomycin, mitotane, mitoxantrone, nifedipine, lutamide, octreotide, oxaliplatin, pamidronate, pentostatin, plicamycin, porfimer, procarbazine, raltitrexed, rituximab, streptozocin, teniposide, testosterone, thalidomide, thioguanine, thiotepa, tretinoin, vindesine, 13-cis retinoic acid, phenylalanine mustard, uracil mustard, estramustine, altretamine, floxuridine, 5-deoxyuridine, cytosine arabinoside, 6-methyl- Lecaptopurine, deoxycoformycin, calcitriol, valrubicin, mithramycin, vinblastine, vinorelbine, topotecan, razoxine, marimastat, COL-3, neovastat, BMS-275291, squalamine, endostatin, SU5416, SU6668, EMD121974, interleukin-12, IM862, angiostatin, vitaxin, droloxifene, idoxifene, spironolactone, finasteride, cimitidine, trase Tuzumab, denileukin diftitox, gefitinib, bortezomib, paclitaxel, cremophor-free paclitaxel, docetaxel, epithilone B, BMS-247550, BMS-310705, droloxifene, 4-hydroxytamoxifen, pipendoxifene, ERA-923, arzoxifene, fulvestrant, acolbifene, lasofoxifene, idoxifene, TSE-424, HMR-3339, ZK186619, topotecan, PTK787/ZK 222584, VX-745, PD 184352, rapamycin, 40-O-(2-hydroxyethyl)-rapamycin, temsirolimus, AP-23573, RAD001, ABT-578, BC-210, LY294002, LY292223, LY292696, LY293684, LY293646, wortmannin, ZM336372, L-779,450, PEG-filgrastim, darbepoetin, erythropoietin, granulocyte colony-stimulating factor, zoledronate , prednisolone, cetuximab, granulocyte-monocyte colony-stimulating factor, histrelin, pegylated interferon alpha-2a, interferon alpha-2a, pegylated interferon alpha-2b, interferon alpha-2b, azacytidine, PEG-L-asparaginase, lenalidomide, gemtuzumab, hydrocortisone, interleukin-11, dexrazoxane, alemtuzumab, all-trans retinoic acid, ketoconazole , interleukin-2, megestrol, immunoglobulin, nitrogen mustard, methylprednisolone, ibritumomab tiuxetan, androgens, decitabine, hexamethylmelamine, bexarotene, tositumomab, arsenic trioxide, cortisone, etidronate, mitotane, cyclosporine, liposomal daunorubicin, edwina-asparaginase, strontium-89, casopitant, netupitant, NK-1 receptor antagonist, paclitaxel These include lonosetron, aprepitant, diphenhydramine, hydroxyzine, metoclopramide, lorazepam, alprazolam, haloperidol, droperidol, dronabinol, dexamethasone, methylprednisolone, prochlorperazine, granisetron, ondansetron, dolasetron, tropisetron, pegfilgrastim, erythropoietin, epoetin alfa, darbepoetin alfa, and mixtures thereof.
「抗HIV剤」または「追加的抗HIV剤」という用語は、例えば特にヌクレオチド逆転写酵素阻害剤(NRTI)、他の非ヌクレオシド逆転写阻害剤(すなわち、本開示の代表的なものではない剤)、プロテアーゼ阻害剤、融合阻害剤を含み、それらの例示的な化合物としては例えば、3TC(ラミブジン(Lamivudine))、AZT(ジドブジン(Zidovudine))、(-)-FTC、ddI(ジダノシン(Didanosine))、ddC(ザルシタビン(zalcitabine))、アバカビル(abacavir)(ABC)、テノホビル(tenofovir)(PMPA)、D-D4FC(レベルセット(Reverset))、D4T(スタブジン(Stavudine))、ラシビル(Racivir)、L-FddC、L-FD4C、NVP(ネビラピン(Nevirapine))、DLV(デラビルジン(Delavirdine))、EFV(えエファビレンツ(Efavirenz))、SQVM(サキナビルメシル酸塩(Saquinavir mesylate))、RTV(リトナビル(Ritonavir))、IDV(インジナビル(Indinavir))、SQV(サキナビル(Saquinavir))、NFV(ネルフィナビル(Nelfinavir))、APV(アムプレナビル(Amprenavir))、LPV(ロピナビル(Lopinavir))、現在治験中または開発中の抗HIV化合物を含む、それらの融合および混合物になどのT20のような融合阻害剤が挙げられる。 The term "anti-HIV agent" or "additional anti-HIV agent" includes, among others, nucleoside reverse transcriptase inhibitors (NRTIs), other non-nucleoside reverse transcriptase inhibitors (i.e., agents not representative of the present disclosure), protease inhibitors, and fusion inhibitors, exemplary compounds of which include, for example, 3TC (lamivudine), AZT (zidovudine), (-)-FTC, ddI (didanosine), ddC (zalcitabine), and the like. abacavir (ABC), tenofovir (PMPA), D-D4FC (Reverset), D4T (Stavudine), Racivir, L-FddC, L-FD4C, NVP (Nevirapine), DLV (Delavirdine), EFV (Efavirenz), SQVM (Saquinavir Mesylate) mesylate), RTV (Ritonavir), IDV (Indinavir), SQV (Saquinavir), NFV (Nelfinavir), APV (Amprenavir), LPV (Lopinavir), fusion inhibitors such as T20, etc., including fusions and mixtures thereof, including anti-HIV compounds currently in clinical trials or development.
本開示に従う化合物との同時投与で使用され得る他の抗HIV剤としては、例えば、とりわけ、ネビラピン(BI-R6-587)、デラビルジン(U-90152S/T)、エファビレンツ(DMP-266)、UC-781(N-[4-クロロ-3-(3-メチル-2-ブテニルオキシ)フェニル]-2メチル3-フランカルボチアミド)、エトラビリン(TMC125)、トロビルジン(Ly300046.HCl)、MKC-442(エミビリン、コアクチノン)、HI-236、HI-240、HI-280、HI-281、リルピビリン(TMC-278)、MSC-127、HBY 097、DMP266、バイカリン(TJN-151)ADAM-II(メチル3’,3’-ジクロロ-4’,4’’-ジメトキシ-5’,5”-ビス(メトキシカルボニル)-6,6-ジフェニルヘキセノエート)、メチル3-ブロモ-5-(1-5-ブロモ-4-メトキシ-3-(メトキシカルボニル)フェニル)フプト-1-エニル)-2-メトキシベンゾエート(アルケニルジアリールメタン類似体、Adam類似体)、(5-クロロ-3-(フェニルスルフィニル)-2’-インドールカルボキサミド)、AAP-BHAP(U-104489またはPNU-104489)、カプラビリン(AG-1549、S-1153)、アテビルジン(U-87201E)、アウリントリカルボン酸(SD-095345)、1-[(6-シアノ-2-インドリル)カルボニル]-4-[3-(イソプロピルアミノ)-2-ピリジニル]ピペラジン、1-[5-[[N-(メチル)メチルスルホニルアミノ]-2-インドリルカルボニル-4-[3-(イソプロピルアミノ)-2-ピリジニル]ピペラジン、1-[3-(エチルアミノ)-2-[ピリジニル]-4-[(5-ヒドロキシ-2-インドリル)カルボニル]ピペラジン、1-[(6-ホルミル-2-インドリル)カルボニル]-4-[3-(イソプロピルアミノ)-2-ピリジニル]ピペラジン、1-[[5-(メチルスルホニルオキシ)-2-インドイリ)カルボニル]-4-[3-(イソプロピルアミノ)-2-ピリジニル]ピペラジン、U88204E、ビス(2-ニトロフェニル)スルホン(NSC633001)、カラノライドA(NSC675451)、カラノライドB、6-ベンジル-5-メチル-2-(シクロヘキシルオキシ)ピリミジン-4-オン(DABO-546)、DPC961、E-EBU、E-EBU-dm、E-EPSeU、E-EPU、フォスカルネット(フォスカルビル)、HEPT(1-[(2-ヒドロキシエトキシ)メチル]-6-(フェニルチオ)チミン)、HEPT-M(1-[(2-ヒドロキシエトキシ)メチル]-6-(3-メチルフェニル)チオ)チミン)、HEPT-S(1-[(2-ヒドロキシエトキシ)メチル]-6-(フェニルチオ)-2-チオチミン)、イノフィラムP、L-737,126、ミケラミンA(NSC650898)、ミケラミンB(NSC649324)、ミケラミンF、6-(3,5-ジメチルベンジル)-1-[(2-ヒドロキシエトキシ)メチル]-5-イソプロピルウラシル、6-(3,5-ジメチルベンジル)-1-(エチオキシメチル)-5-イソプロピルウラシル、NPPS、E-BPTU(NSC648400)、オルチプラズ(4-メチル-5-(ピラジニル)-3H-1,2-ジチオール-3-チオン)、N-{2-(2-クロロ-6-フルオロフェネチル]-N’-(2-チアゾリル)チオ尿素(PETT Cl、F誘導体)、N-{2-(2,6-ジフルオロフェネチル]-N’-[2-(5-ブロモピリジル)]チオ尿素{PETT誘導体)、N-{2-(2,6-ジフルオロフェネチル]-N’-[2-(5-メチルピリジル)]チオ尿素{PETTピリジル誘導体)、N-[2-(3-フルオロフラニル)エチル]-N’-[2-(5-クロロピリジル)]チオ尿素、N-[2-(2-フルオロ-6-エトキシフェネチル)]-N’-[2-(5-ブロモピリジル)]チオ尿素、N-(2-フェネチル)-N’-(2-チアゾリル)チオ尿素(LY-73497)、L-697,639、L-697,593、L-697,661、3-[2-(4,7-ジフルオロベンゾオキサゾール-2-イル)エチル}-5-エチル-6-メチル(ピプリジン-2(1H)-チオン(2-ピリジノン誘導体)、3-[[(2-メトキシ-5,6-ジメチル-3-ピリジル)メチル]アミン]-5-エチル-6-メチル(ピプリジン-2(1H)-チオン、R82150、R82913、R87232、R88703、R89439(ロビリド)、R90385、S-2720、スラミンナトリウム、TBZ(チアゾロベンズイミダゾール、NSC 625487)、チアゾロイソインドール-5-オン、(+)(R)-9b-(3,5-ジメチルフェニル-2,3-ジヒドロチアゾロ[2,3-a]イソインドール-5(9bH)-オン、チビラピン(R86183)、UC-38、およびUC-84からなる群から選択され得る他のNNRTI(すなわち、本開示に従うNNRTI以外)が挙げられる。 Other anti-HIV agents that may be used in co-administration with the compounds according to the present disclosure include, for example, nevirapine (BI-R6-587), delavirdine (U-90152S/T), efavirenz (DMP-266), UC-781 (N-[4-chloro-3-(3-methyl-2-butenyloxy)phenyl]-2-methyl-3-furancarbothiamide), etravirine (TMC125), trovirdine (Ly300046.HCl), MKC-442 (emivirine, coactinone), HI-236, HI-240, HI-280, HI-281, rilpivirine (TMC-278), MSC-127, HBY, among others. 097, DMP266, baicalin (TJN-151) ADAM-II (methyl 3',3'-dichloro-4',4''-dimethoxy-5',5"-bis(methoxycarbonyl)-6,6-diphenylhexenoate), methyl 3-bromo-5-(1-5-bromo-4-methoxy-3-(methoxycarbonyl)phenyl)hup-1-enyl)-2-methoxybenzoate (alkenyl diarylmethane analog, Adam analog), (5-chloro-3-(phenylsulfinyl)-2'-indolecarboxamide), AAP-BHAP (U-104489 or PNU-104489), capravirine (AG-1549, S-1153), atevirdine (U-87201 E), aurintricarboxylic acid (SD-095345), 1-[(6-cyano-2-indolyl)carbonyl]-4-[3-(isopropylamino)-2-pyridinyl]piperazine, 1-[5-[[N-(methyl)methylsulfonylamino]-2-indolylcarbonyl-4-[3-(isopropylamino)-2-pyridinyl]piperazine, 1-[3-(ethylamino)-2-[pyridinyl]-4-[(5-hydroxy-2-indolyl)carbonyl]piperazine, 1-[(6-formyl-2-indolyl)carbonyl]-4-[3-(isopropylamino)-2-pyridinyl]piperazine, 1-[[5-(methylsulfonyloxy)-2-indolyl)carbonyl]- 4-[3-(isopropylamino)-2-pyridinyl]piperazine, U88204E, bis(2-nitrophenyl)sulfone (NSC 633001), calanolide A (NSC 675451), calanolide B, 6-benzyl-5-methyl-2-(cyclohexyloxy)pyrimidin-4-one (DABO-546), DPC961, E-EBU, E-EBU-dm, E-EPSeU, E-EPU, foscarnet (foscarbil), HEPT (1-[(2-hydroxyethoxy)methyl]-6-(phenylthio)thymine), HEPT-M (1-[(2-hydroxyethoxy)methyl]-6-(3-methylphenyl)thio)thymine), HEPT-S (1-[(2-hydroxyethoxy)methyl]-6-(phenylthio)thymine), 6-(3,5-dimethylbenzyl)-1-[(2-hydroxyethoxy)methyl]-5-isopropyluracil, 6-(3,5-dimethylbenzyl)-1-(ethyoxymethyl)-5-isopropyluracil, NPPS, E-BPTU (NSC 648400), Oltipraz (4-methyl-5-(pyrazinyl)-3H-1,2-dithiole-3-thione), N-{2-(2-chloro-6-fluorophenethyl]-N'-(2-thiazolyl)thiourea (PETT Cl, F derivatives), N-{2-(2,6-difluorophenethyl]-N'-[2-(5-bromopyridyl)]thiourea {PETT derivatives), N-{2-(2,6-difluorophenethyl]-N'-[2-(5-methylpyridyl)]thiourea {PETT pyridyl derivatives), N-[2-(3-fluorofuranyl)ethyl]-N'-[2-(5-chloropyridyl)]thiourea, N-[2-(2-fluoro-6-ethoxyphenethyl)]-N'-[2-(5-bromopyridyl)]thiourea, N-(2-phenethyl)-N'-(2-thiazolyl)thiourea (LY-73497), L-697,639, L-697,593, L-697,661, 3-[2-(4,7-difluorobenzoxazol-2-yl)ethyl}-5-ethyl-6-methyl(pipridine-2(1H)-thione (2-pyridinone derivative), 3-[[(2-methoxy-5,6-dimethyl-3-pyridyl)methyl]amine]-5-ethyl-6-methyl(pipridine-2(1H)-thione, R82150, R82913, R87232, R88703, R89439 (Roviride), R90385, S-2720, Suramin sodium, TBZ (thiazolbenzimidazole, NSC 625487), thiazoloisoindol-5-one, (+)(R)-9b-(3,5-dimethylphenyl-2,3-dihydrothiazolo[2,3-a]isoindol-5(9bH)-one, tivirapine (R86183), UC-38, and UC-84.
「薬学的に許容可能な塩」という用語は、本明細書全体を通して適切な場合に本明細書に記載の化合物のうちの1つ以上の塩形態を記載するために使用され、それら塩形態は、患者の消化管の胃液において化合物の可溶性増加を示し、当該化合物の溶解および生体利用効率を促進する。薬学的に許容可能な塩としては、適切な場合には薬学的に許容可能な無機または有機の塩基および酸から誘導されるものが挙げられる。適切な塩としては特に薬学分野において公知の多くの他の酸および塩基のなかで、例えばカリウムおよびナトリウムなどのアルカリ金属、例えばカルシウム、マグネシウムなどのアルカリ土類金属、およびアンモニウム塩に由来するものが挙げられる。ナトリウムおよびカリウムの塩は、本開示によるリン酸塩の中和塩として特に好ましい。 The term "pharmaceutically acceptable salt" is used throughout the specification to describe, where appropriate, one or more salt forms of the compounds described herein that exhibit increased solubility of the compound in the gastric fluids of the patient's digestive tract and promote dissolution and bioavailability of the compound. Pharmaceutically acceptable salts include those derived from pharmaceutically acceptable inorganic or organic bases and acids, where appropriate. Suitable salts include those derived from alkali metals, e.g., potassium and sodium, alkaline earth metals, e.g., calcium, magnesium, and ammonium salts, among many other acids and bases known in the pharmaceutical art. Sodium and potassium salts are particularly preferred as neutralizing salts of phosphates according to the present disclosure.
「薬剤的に許容される誘導体」という語は、本明細書全体を通じて任意の薬剤的に許容されるプロドラッグ形態(例えばエステル、アミド、他のプロドラッグ群など)を記述するために使用され、それらは患者に投与すると本化合物を直接または間接的に生じさせ、または本化合物の活性代謝物を生じさせる。 The term "pharmaceutical acceptable derivative" is used throughout this specification to describe any pharmaceutical acceptable prodrug form (e.g., esters, amides, other prodrug groups, etc.) that upon administration to a patient, directly or indirectly yields the compound or an active metabolite of the compound.
一般的な合成アプローチ
本明細書に記載される二官能性分子の合成の実現および最適化は、段階的またはモジュール形式で取り組んでもよい。例えば、標的分子に結合する化合物の特定は、適切なリガンドが直ちに利用できない場合、ハイスループット、またはミディアムスループットのスクリーニングキャンペーンを含んでもよい。適切なインビトロの薬理アッセイおよび/またはADMETアッセイのデータにより特定される次善の態様を改善するために、初期リガンドに対し、反復設計およびサイクル最適化が必要とされることは珍しいことではない。最適化/SARキャンペーンの一部は、置換に忍容性があり、本明細書において上記に参照されたリンカー化学物質を結合させる適切な位置であり得るリガンド位置をプローブすることである。結晶学的またはNMR構造データが利用可能である場合、これらを使用して、かかる合成的な取り組みに焦点を当てることができる。
General synthetic approach The realization and optimization of the synthesis of the bifunctional molecules described herein may be undertaken in a stepwise or modular fashion. For example, the identification of compounds that bind to a target molecule may involve high-throughput or medium-throughput screening campaigns when suitable ligands are not readily available. It is not uncommon that iterative design and cycle optimization is required for initial ligands to improve suboptimal aspects identified by suitable in vitro pharmacological and/or ADMET assay data. Part of the optimization/SAR campaign is to probe ligand positions that tolerate substitution and may be suitable positions for attaching the linker chemistry referenced herein above. When crystallographic or NMR structural data are available, these can be used to focus such synthetic efforts.
非常に類似した方法で、E3リガーゼ、すなわち、ULM/VLMのリガンドを特定し、最適化することができる。 In a very similar manner, ligands for E3 ligases, i.e., ULM/VLM, can be identified and optimized.
PTMおよびULM(例えば、VLMs)が手元にある場合、当業者であれば、リンカー部分の有無にかかわらず、それらの組み合わせのために既知の合成方法を使用することができる。PTM基およびULM基がリンカーの遠位端に連続して結合され得るように、リンカー部分をある範囲の組成、長さ、および柔軟性で合成し、官能基化することができる。したがって、インビトロおよびインビボの薬理学的試験、およびADMET/PK試験において、二官能性分子のライブラリを実現化し、プロファイリングすることができる。PTM基およびULM基と同様に、最終的な二官能性分子に反復設計を施し、最適化のサイクルを行って、所望の特性を有する分子を特定することができる。 With PTMs and ULMs (e.g., VLMs) at hand, one of skill in the art can use known synthetic methods for their combination with or without linker moieties. Linker moieties can be synthesized and functionalized with a range of compositions, lengths, and flexibility so that PTM and ULM groups can be successively attached to the distal ends of the linker. Thus, libraries of bifunctional molecules can be realized and profiled in in vitro and in vivo pharmacological and ADMET/PK studies. As with the PTM and ULM groups, the final bifunctional molecules can be subjected to cycles of iterative design and optimization to identify molecules with desired properties.
一部の例では、保護基戦略および/または官能基相互変換(FGI)が、所望材料の調製を促進するために必要とされる場合がある。そのような化学的プロセスは合成有機化学者に公知であり、これら化学的プロセスの多くが、例えば“Greene’s Protective Groups in Organic Synthesis”Peter G.M.Wuts and Theodora W.Greene(Wiley)、および“Organic Synthesis:The Disconnection Approach”Stuart Warren and Paul Wyatt(Wiley)などの書籍に見出され得る。 In some cases, protecting group strategies and/or functional group interconversions (FGI) may be required to facilitate the preparation of the desired material. Such chemical processes are known to synthetic organic chemists, and many of these chemical processes can be found in books such as, for example, "Greene's Protective Groups in Organic Synthesis" by Peter G. M. Wuts and Theodora W. Greene (Wiley), and "Organic Synthesis: The Disconnection Approach" by Stuart Warren and Paul Wyatt (Wiley).
略語:
ACN:アセトニトリル
ADDP:1,1’-(アゾジカルボニル)ジピペリジン
BAST:N,N-ビス(2-メトキシエチル)アミノ硫黄トリフルオリド
BPO:過酸化ベンゾイル
Cbz:カルボニルベジルオキシ
DAST:ジエチルアミノ硫黄トリフルオリド
DBE:1,2-ジブロモエタン
DCM:ジクロロメタン
DEAD:アゾジカルボン酸ジエチル
DIAD:アゾジプロピル酸ジイソプロピル
DIBAL:水素化ジシオブチルアルミニウム
DIEAまたはDIPEA:ジイソプロピルエチルアミン
DMA:N,N-ジメチルアセトアミド
DMF:N,N-ジメチルホルムアミド
DMP:デス-マーチンペルヨージナン
EA:酢酸エチル
EDCI:1-エチル-3-(3-ジメチルアミノプロピル)カルボジイミド
HBTU:N,N,N’N’-テトラメチル-O-(1H-ベンゾトリアゾール-1-イル)ウロニウムヘキサフルオロリン酸塩
HMDS:ビス(トリメチルシリル)アミン
HMPA:ヘキサメチルホスホラミド
LDA:リチウムジイソプロピルアミド
MCPBA:メタ-クロロ過安息香酸
MsCl:塩化メタンスルホニル
M.W:マイクロ波
NBS:N-ブロモスクシンイミド
NMP:N-メチルピロリドン
PCC:クロロクロム酸ピリジニウム
Pd-118またはPd(dtpf)Cl2:1,1’-ビス(ジ-tert-ブチルホスフィノ)フェロセンジクロロパラジウム
Pd(dppf)Cl2:1,1’-ビス(ジフェニルホスフィノ)フェロセンジクロロパラジウム
Pd(dba)2:ビス(ジベンジリデンアセトン)パラジウム
Pd2(dba)3:トリス(ジベンジリデンアセトン)ジパラジウム
PPTS:p-トルンスルホン酸ピリジウム
PTSA:p-トルエンスルホン酸
RuPhos-Pd-G3:XPhos-Pd-G3:[(2-ジシクロヘキシルホスフィノ-2′,6′-ジイソプロポキシ-1,1′-ビフェニル)-2-(2′-アミノ-1,1′-ビフェニル)]パラジウム(II)メタンスルホネート
RuPhos-Pd-G2:クロロ[(2-ジシクロヘキシルホスフィノ-2’,6’-ジイソプロポキシ-1,1’-ビフェニル)-2-(2’-アミノ-1,1’-ビフェニル)]パラジウム(II)
SFC:超臨界流体クロマトグラフィー
t-BuXPhos-Pd-G3:[(2-ジ-tert-ブチルホスフィノ-2′,4′,6′-トリイソプロピル-1,1′-ビフェニル)-2-(2′-アミノ-1,1′-ビフェニル)]パラジウム(II)メタンスルホネート
TEA:トリメチルアミン
TFA:トリフルオロ酢酸
TLC:薄層クロマトグラフィー
TMP:2,2,6,6-テトラメチルピペリジン
TEMPO:2,2,6,6-テトラメチルピペリジン-N-オキシド
TosClまたはTsCl:p-トルエンスルホニルクロリド
TsOH:p-トルエンスルホン酸
XantPhos:4,5-ビス(ジフェニルホスフィノ)-9,9-ジメチルキサンテン
XPhos:2-ジクロヘキシルホスフィノ-2’4’6’-トリイソプロピルビフェニル
XPhos-Pd-G3:[(2-ジシクロヘキシルホスフィノ-2′,4′,6′-トリイソプロピル-1,1′-ビフェニル)-2-(2′-アミノ-1,1′-ビフェニル)]パラジウム(II)メタンスルホネート
12354-85-7:ビス(ペンタメチルシクロペンタジエニルロジウム二塩化物)
Abbreviations:
ACN: Acetonitrile ADDP: 1,1'-(azodicarbonyl)dipiperidine BAST: N,N-bis(2-methoxyethyl)aminosulfur trifluoride BPO: Benzoyl peroxide Cbz: Carbonylbenzyloxy DAST: Diethylaminosulfur trifluoride DBE: 1,2-dibromoethane DCM: Dichloromethane DEAD: Diethyl azodicarboxylate DIAD: Diisopropyl azodipropylate DIBAL: Dithiobutylaluminum hydride DIEA or DIPEA: Diisopropylethylamine DMA: N,N-Dimethylacetamide DMF: N,N-Dimethylformamide DMP: Dess-Martin periodinane EA: Ethyl acetate EDCI: 1-Ethyl-3-(3-dimethylaminopropyl)carbodiimide HBTU: N,N,N'N'-tetramethyl-O-(1H-benzotriazol-1-yl)uronium hexafluorophosphate HMDS: bis(trimethylsilyl)amine HMPA: hexamethylphosphoramide LDA: lithium diisopropylamide MCPBA: meta-chloroperbenzoic acid MsCl: methanesulfonyl chloride M. W: Microwave NBS: N-bromosuccinimide NMP: N-methylpyrrolidone PCC: Pyridinium chlorochromate Pd-118 or Pd(dtpf)Cl2: 1,1'-bis(di-tert-butylphosphino)ferrocenedichloropalladium Pd(dppf)Cl2: 1,1'-bis(diphenylphosphino)ferrocenedichloropalladium Pd(dba)2: Bis(dibenzylideneacetone)palladium Pd2(dba)3: Tris(dibenzylideneacetone)dipalladium PPTS: Pyridinium p-toluenesulfonate PTSA: p-toluenesulfonic acid RuPhos-Pd-G3: XPhos-Pd-G3: [(2-dicyclohexylphosphino-2',6'-diisopropoxy-1,1'-biphenyl)-2-(2'-amino-1,1'-biphenyl)]palladium(II) methanesulfonate RuPhos-Pd-G2: Chloro[(2-dicyclohexylphosphino-2',6'-diisopropoxy-1,1'-biphenyl)-2-(2'-amino-1,1'-biphenyl)]palladium(II)
SFC: Supercritical fluid chromatography t-BuXPhos-Pd-G3: [(2-di-tert-butylphosphino-2',4',6'-triisopropyl-1,1'-biphenyl)-2-(2'-amino-1,1'-biphenyl)]palladium(II) methanesulfonate TEA: Trimethylamine TFA: Trifluoroacetic acid TLC: Thin layer chromatography TMP: 2,2,6,6-tetramethylpiperidine TEMPO: 2,2,6,6-tetramethylpiperidine-N-oxide TosCl or TsCl: p-toluenesulfonyl chloride TsOH: p-toluenesulfonic acid XantPhos: 4,5-bis(diphenylphosphino)-9,9-dimethylxanthene XPhos: 2-dicyclohexylphosphino-2'4'6'-triisopropylbiphenyl XPhos-Pd-G3: [(2-dicyclohexylphosphino-2',4',6'-triisopropyl-1,1'-biphenyl)-2-(2'-amino-1,1'-biphenyl)]palladium(II) methanesulfonate 12354-85-7: Bis(pentamethylcyclopentadienylrhodium dichloride)
スキーム1

スキーム1に示されるように、WPTM5部分を有する化合物は、アミノ基などの求核基を含む結合基Lを含有する。良好な脱離基LGを有する化合物(例えば、ペルフルオロスルホニル基C4F9SO3-)と反応させると、カップリング生成物が形成される。カップリング生成物は、パラジウム触媒化条件下で単保護アミン分子WPTM3と反応して、WPTM3断片に結合する。アミン脱保護に続いて、WPTM5-WPTM3中のZ’およびZ’’から選択されるより反応性の高いハロゲン原子を、遊離アミノ基で求核置換して、WPTM5-WPTM3-WPTM2を形成し、次いで、WPTM2ボロン酸と反応させて、WPTM5-WPTM3-WPTM2-WPTM1断片を含むPTM結合基を提供する。塩基触媒化エステル加水分解後、得られた酸を、アミノ基を有するULM部分と結合させて、PTMおよびULM結合部分を1つの分子に組み合わせる。 As shown in Scheme 1, compounds bearing a WPTM5 moiety contain a linking group L that includes a nucleophilic group such as an amino group. When reacted with a compound that has a good leaving group LG (e.g., the perfluorosulfonyl group C4F9SO3-), a coupling product is formed. The coupling product is reacted with a monoprotected amine molecule WPTM3 under palladium catalyzed conditions to couple to the WPTM3 fragment. Following amine deprotection, the more reactive halogen atom selected from Z' and Z'' in WPTM5-WPTM3 is nucleophilically displaced with the free amino group to form WPTM5-WPTM3-WPTM2, which is then reacted with WPTM2 boronic acid to provide a PTM linking group that includes the WPTM5-WPTM3-WPTM2-WPTM1 fragment. After base catalyzed ester hydrolysis, the resulting acid is coupled with a ULM moiety bearing an amino group to combine the PTM and ULM linking moieties into one molecule.
スキーム2

スキーム2に示されるように、ヒドロキシル含有部分R3と、結合基Lを含む単保護ジオールとの間のミツノブ反応により、カップリング生成物を得る。O脱保護に続いて、遊離ヒドロキシル基を、例えば、スルホネートとして活性化し、その後、WPTM5においてアミノ基を有する部分WPTM5-WPTM3-WPTM2-WPTM1で置換する。エステル加水分解後、得られた酸を、アミノ基を有するULM部分と結合させて、PTMおよびULM結合部分を1つの分子に組み合わせる。 As shown in Scheme 2, a Mitsunobu reaction between a hydroxyl-containing moiety R3 and a monoprotected diol containing a linking group L gives the coupling product. Following O-deprotection, the free hydroxyl group is activated, for example as a sulfonate, and then substituted with a moiety bearing an amino group in WPTM5, WPTM5-WPTM3-WPTM2-WPTM1. After ester hydrolysis, the resulting acid is coupled with a ULM moiety bearing an amino group to combine the PTM and ULM linking moieties into one molecule.
スキーム3

スキーム3に示されるように、ビニル基を、最初に、ビス-ハロゲン化誘導体WPTM2に導入する。次いで、得られた生成物は、任意に保護されたアミノ基を含む結合基L’を有するハロゲン化部分WPTM2とのパラジウム触媒化Heckカップリング反応を受ける。WPTM1部分を導入するために、WPTM2-WPTM5のハロゲン化物部分を、スズキ条件下で適切なボロン酸と結合させ、得られたアミンを、ULM含有アルデヒドと反応させて、PTM-ULMカップリング生成物をもたらす。 As shown in Scheme 3, a vinyl group is first introduced into the bis-halogenated derivative WPTM2. The resulting product then undergoes a palladium-catalyzed Heck coupling reaction with a halogenated moiety WPTM2 bearing a linking group L' containing an optionally protected amino group. To introduce the WPTM1 moiety, the halide moiety of WPTM2-WPTM5 is coupled with an appropriate boronic acid under Suzuki conditions, and the resulting amine is reacted with a ULM-containing aldehyde to yield the PTM-ULM coupling product.
スキーム4

スキーム4は、PTM結合部分WPTM5-WPTM3-WPTM2-WPTM1をULM含有部分と連結するために利用される例示的なカップリング反応を例示する。このような反応は、還元剤としてシアノ水素化ホウ素ナトリウムを使用した還元的アミノ化、またはカルボン酸とジアミンとの間の縮合カップリング反応を含み得る。当業者であれば、適切な試薬および条件を選択して、所望の変換を行うことが可能であろう。 Scheme 4 illustrates exemplary coupling reactions utilized to link the PTM binding moieties WPTM5-WPTM3-WPTM2-WPTM1 with ULM-containing moieties. Such reactions may include reductive amination using sodium cyanoborohydride as a reducing agent, or condensation coupling reactions between carboxylic acids and diamines. One of skill in the art will be able to select appropriate reagents and conditions to effect the desired transformation.
スキーム5

スキーム5に示されるように、ヒドロキシル含有部分R3とWPTM5との間のミツノブ型反応により、カップリング生成物を得る。次いで、WPTM5におけるハロゲン原子を、スズキまたはバックワルド条件下で、単保護ジアミンWPTM3で置換する。第2のアミノ基の脱保護に続いて、WPTM2部分に結合したより反応性の高いハロゲン原子を置換して、WPTM5-WPTM3-WPTM2断片を形成する。次いで、得られたモノハロゲン化物を、スズキ条件下で適切なボロン酸と反応させて、PTM結合部分WPTM5-WPTM3-WPTM2-WPTM2を得る。エステル加水分解後、次いで、得られた酸を、アミノ基を有するULM部分と結合させて、PTMおよびULM結合部分を1つの分子に組み合わせる。 As shown in Scheme 5, a Mitsunobu-type reaction between the hydroxyl-containing moiety R3 and WPTM5 gives the coupling product. The halogen atom in WPTM5 is then displaced with the monoprotected diamine WPTM3 under Suzuki or Buchwald conditions. Following deprotection of the second amino group, the more reactive halogen atom attached to the WPTM2 moiety is displaced to form the WPTM5-WPTM3-WPTM2 fragment. The resulting monohalide is then reacted with an appropriate boronic acid under Suzuki conditions to give the PTM-linked moiety WPTM5-WPTM3-WPTM2-WPTM2. After ester hydrolysis, the resulting acid is then coupled with the ULM moiety bearing an amino group to combine the PTM and ULM-linked moieties into one molecule.
スキーム6

スキーム6に示されるように、アセタール含有ULM部分は、十分に穏やかな条件下で酸触媒化加水分解を受けて、アルデヒドを形成し得、次いで、これを、還元的アミノ化条件下でアミノ基含有リンカーL’を有するWPTM5-WPTM3-WPTM2-WPTM2断片と反応させて、PTM-ULMカップリング生成物を形成する。当業者であれば、適切な試薬および条件を選択して、所望の変換を行うことが可能であろう。 As shown in Scheme 6, the acetal-containing ULM moiety can undergo acid-catalyzed hydrolysis under sufficiently mild conditions to form an aldehyde, which can then be reacted with the WPTM5-WPTM3-WPTM2-WPTM2 fragment bearing an amino group-containing linker L' under reductive amination conditions to form the PTM-ULM coupling product. One of skill in the art will be able to select appropriate reagents and conditions to effect the desired transformation.
スキーム7

スキーム7に示されるように、ヒドロキシル含有部分R3と単保護フッ素化ジオールとの間のミツノブ型反応により、フッ素含有中間体を提供する。O脱保護に続いて、遊離ヒドロキシル基を活性化し(例えば、スルホネートとして)、その後、WPTM5においてアミノ基を有する部分WPTM5-WPTM3-WPTM2-WPTM1で置換する。エステル加水分解後、得られた酸を、アミノ基を有するULM部分と結合させて、PTMおよびULM結合部分を1つの分子に組み合わせる。 As shown in Scheme 7, a Mitsunobu-type reaction between a hydroxyl-containing moiety R3 and a monoprotected fluorinated diol provides a fluorine-containing intermediate. Following O-deprotection, the free hydroxyl group is activated (e.g., as a sulfonate) and then substituted with an amino-bearing moiety WPTM5-WPTM3-WPTM2-WPTM1 in WPTM5. After ester hydrolysis, the resulting acid is coupled with an amino-bearing ULM moiety to combine the PTM and ULM-linked moieties into one molecule.
スキーム8

スキーム8に示されるように、R3を含むヒドロキシル基含有エステルでの、L’-OLG中の脱離基の求核置換により、リンカーL’およびL’’を組み合わせるカップリング生成物を得る。その後の塩基加水分解、および得られた酸と、アミノ基を有するULM部分とのカップリングにより、2つの連続した結合基L’およびL’’が結合するULM断片を提供する。L’のアミノ基のN脱保護、続いてWPTM5-WPTM3-WPTM2-WPTM1-L’”アルデヒドによる還元的アミノ酸化、またはWPTM5-WPTM3-WPTM2-WPTM1-L’”カルボン酸との縮合により、PTM-ULMカップリング生成物を得る。 As shown in Scheme 8, nucleophilic displacement of the leaving group in L'-OLG with a hydroxyl-containing ester containing R3 gives a coupling product combining linkers L' and L". Subsequent base hydrolysis and coupling of the resulting acid with a ULM moiety bearing an amino group provides a ULM fragment with two consecutive linking groups L' and L". N-deprotection of the amino group of L' followed by reductive amino acid conversion with WPTM5-WPTM3-WPTM2-WPTM1-L'" aldehyde or condensation with WPTM5-WPTM3-WPTM2-WPTM1-L'" carboxylic acid gives the PTM-ULM coupling product.
スキーム9
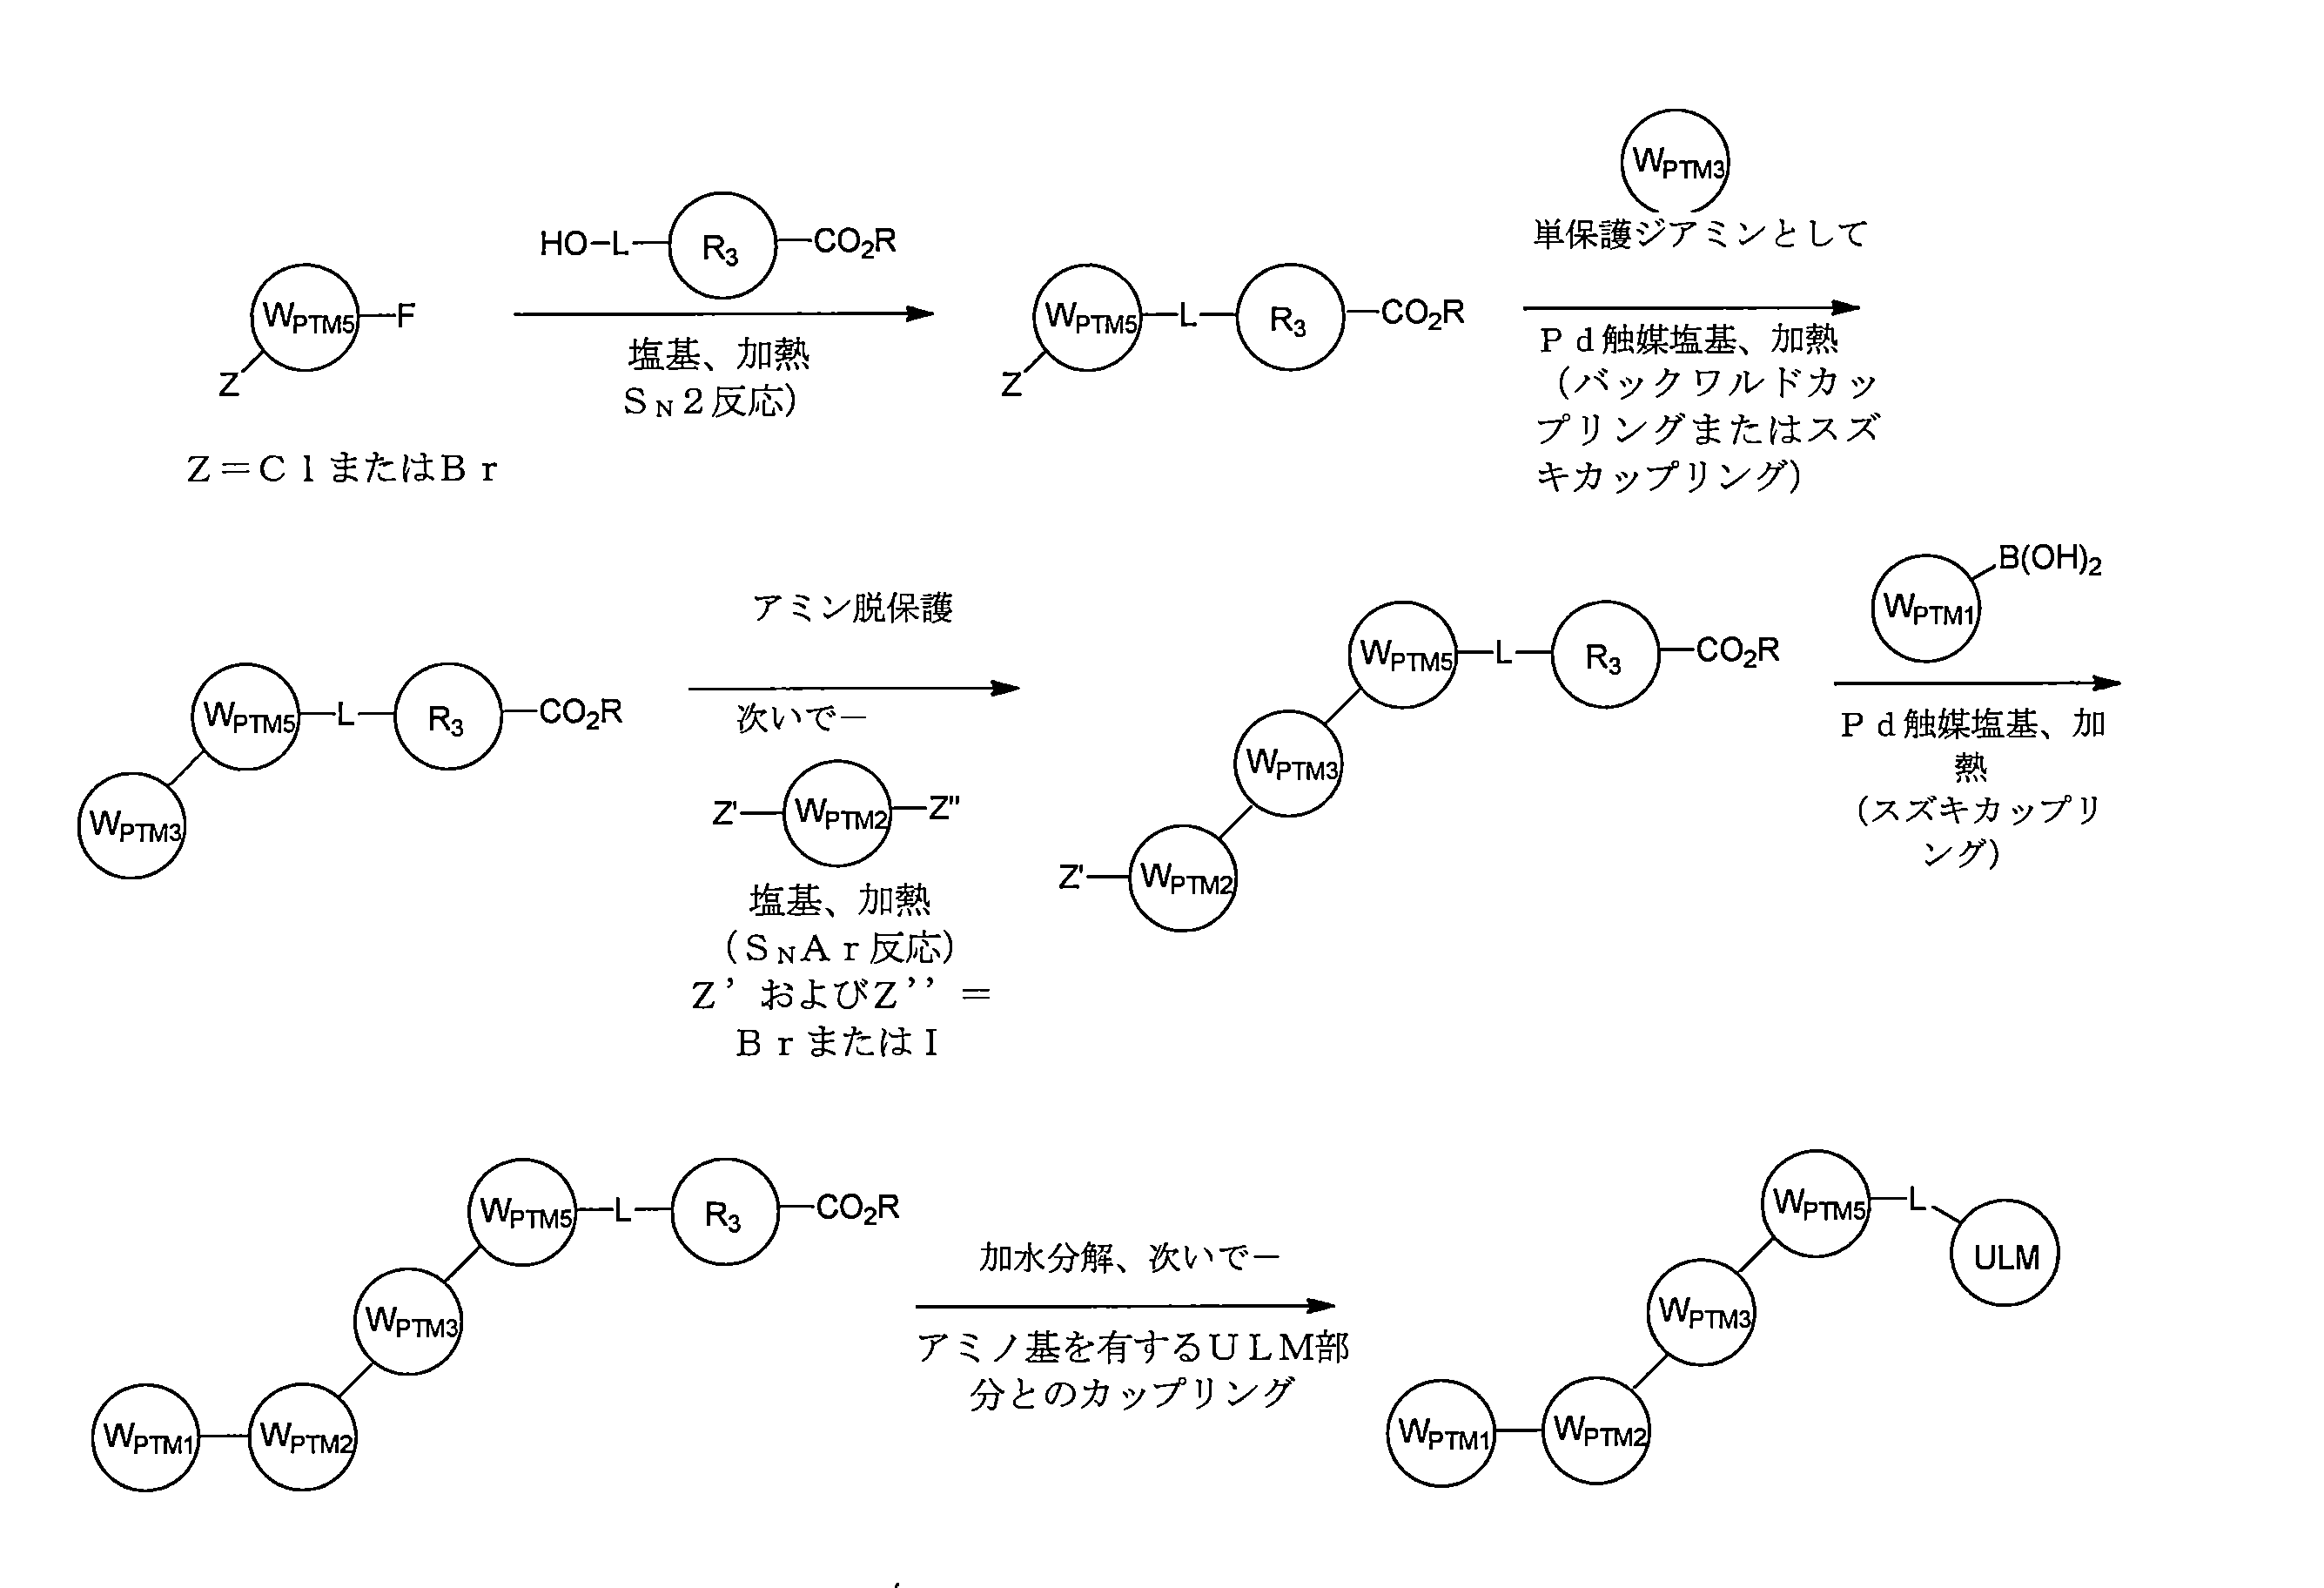
スキーム9に示されるように、R3を有するヒドロキシル基含有化合物での、部分WPTM5中のフッ素原子の求核置換により、WPTM5-L-R3ハロゲン化物を提供し、これを、スズキまたはバックワルド条件下で単保護ジアミンWPTM5に結合させる。第2のアミノ基の脱保護に続いて、次いで、WPTM3-WPTM5-L-R3部分を芳香族求核置換によりビスハロゲン化WPTM2と反応させて、モノハロゲン化物を提供する。その後の適切なボロン酸とのスズキ反応により、WPTM1-WPTM2-WPTM3-WPTM5-L-R3エステルを提供する。塩基加水分解およびアミノ基を有するULM部分とのカップリングにより、PTM-ULMカップリング生成物を提供する。 As shown in Scheme 9, nucleophilic displacement of the fluorine atom in the moiety WPTM5 with a hydroxyl group-containing compound bearing R3 provides the WPTM5-L-R3 halide, which is coupled to the monoprotected diamine WPTM5 under Suzuki or Buchwald conditions. Following deprotection of the second amino group, the WPTM3-WPTM5-L-R3 moiety is then reacted with the bishalogenated WPTM2 by aromatic nucleophilic displacement to provide the monohalide. Subsequent Suzuki reaction with an appropriate boronic acid provides the WPTM1-WPTM2-WPTM3-WPTM5-L-R3 ester. Base hydrolysis and coupling with an amino-bearing ULM moiety provides the PTM-ULM coupling product.
本開示の例示的な化合物を合成するための1つの可能なアプローチは、以下のスキームに詳述される一般的な合成経路に従うものである。

当業者であれば、改変されたアプローチを利用して、異なる化学リンカーを介してPTMを結合することができることを理解するであろう。例えば、WPTM5がCH2基を介してL’に連結されている場合(上記スキームにおいてX=CH2)では、以下のスキームに記載されるアプローチを想定することができる。

あるいは、WPTM5が存在しない場合、式Iによって表される例示的な化合物のPTMは、以下の一般スキームに従って合成することができる。

当業者であれば、本明細書に記載される一般的なアプローチを改変して、WPTM1、WPTM2、WPTM3、WPTM4、WPTM5、WPTM6、およびWPTM7環の特定の性質に適合させることができることを理解するであろう。例えば、いくつかの実施形態では、式IIによって表される例示的な化合物は、当業者であれば、例示的な化合物の特定の化学的性質に応じて、追加の保護/脱保護工程が必要とされ得ることを認識するであろう、以下の一般的な合成スキームに記載されるように調整することができる。

XがNHを表す一実施形態では、例示的な化合物は、WPTM5が存在するかどうかに応じて、以下に示す2つのスキームのうちの1つに従って調製することができる。
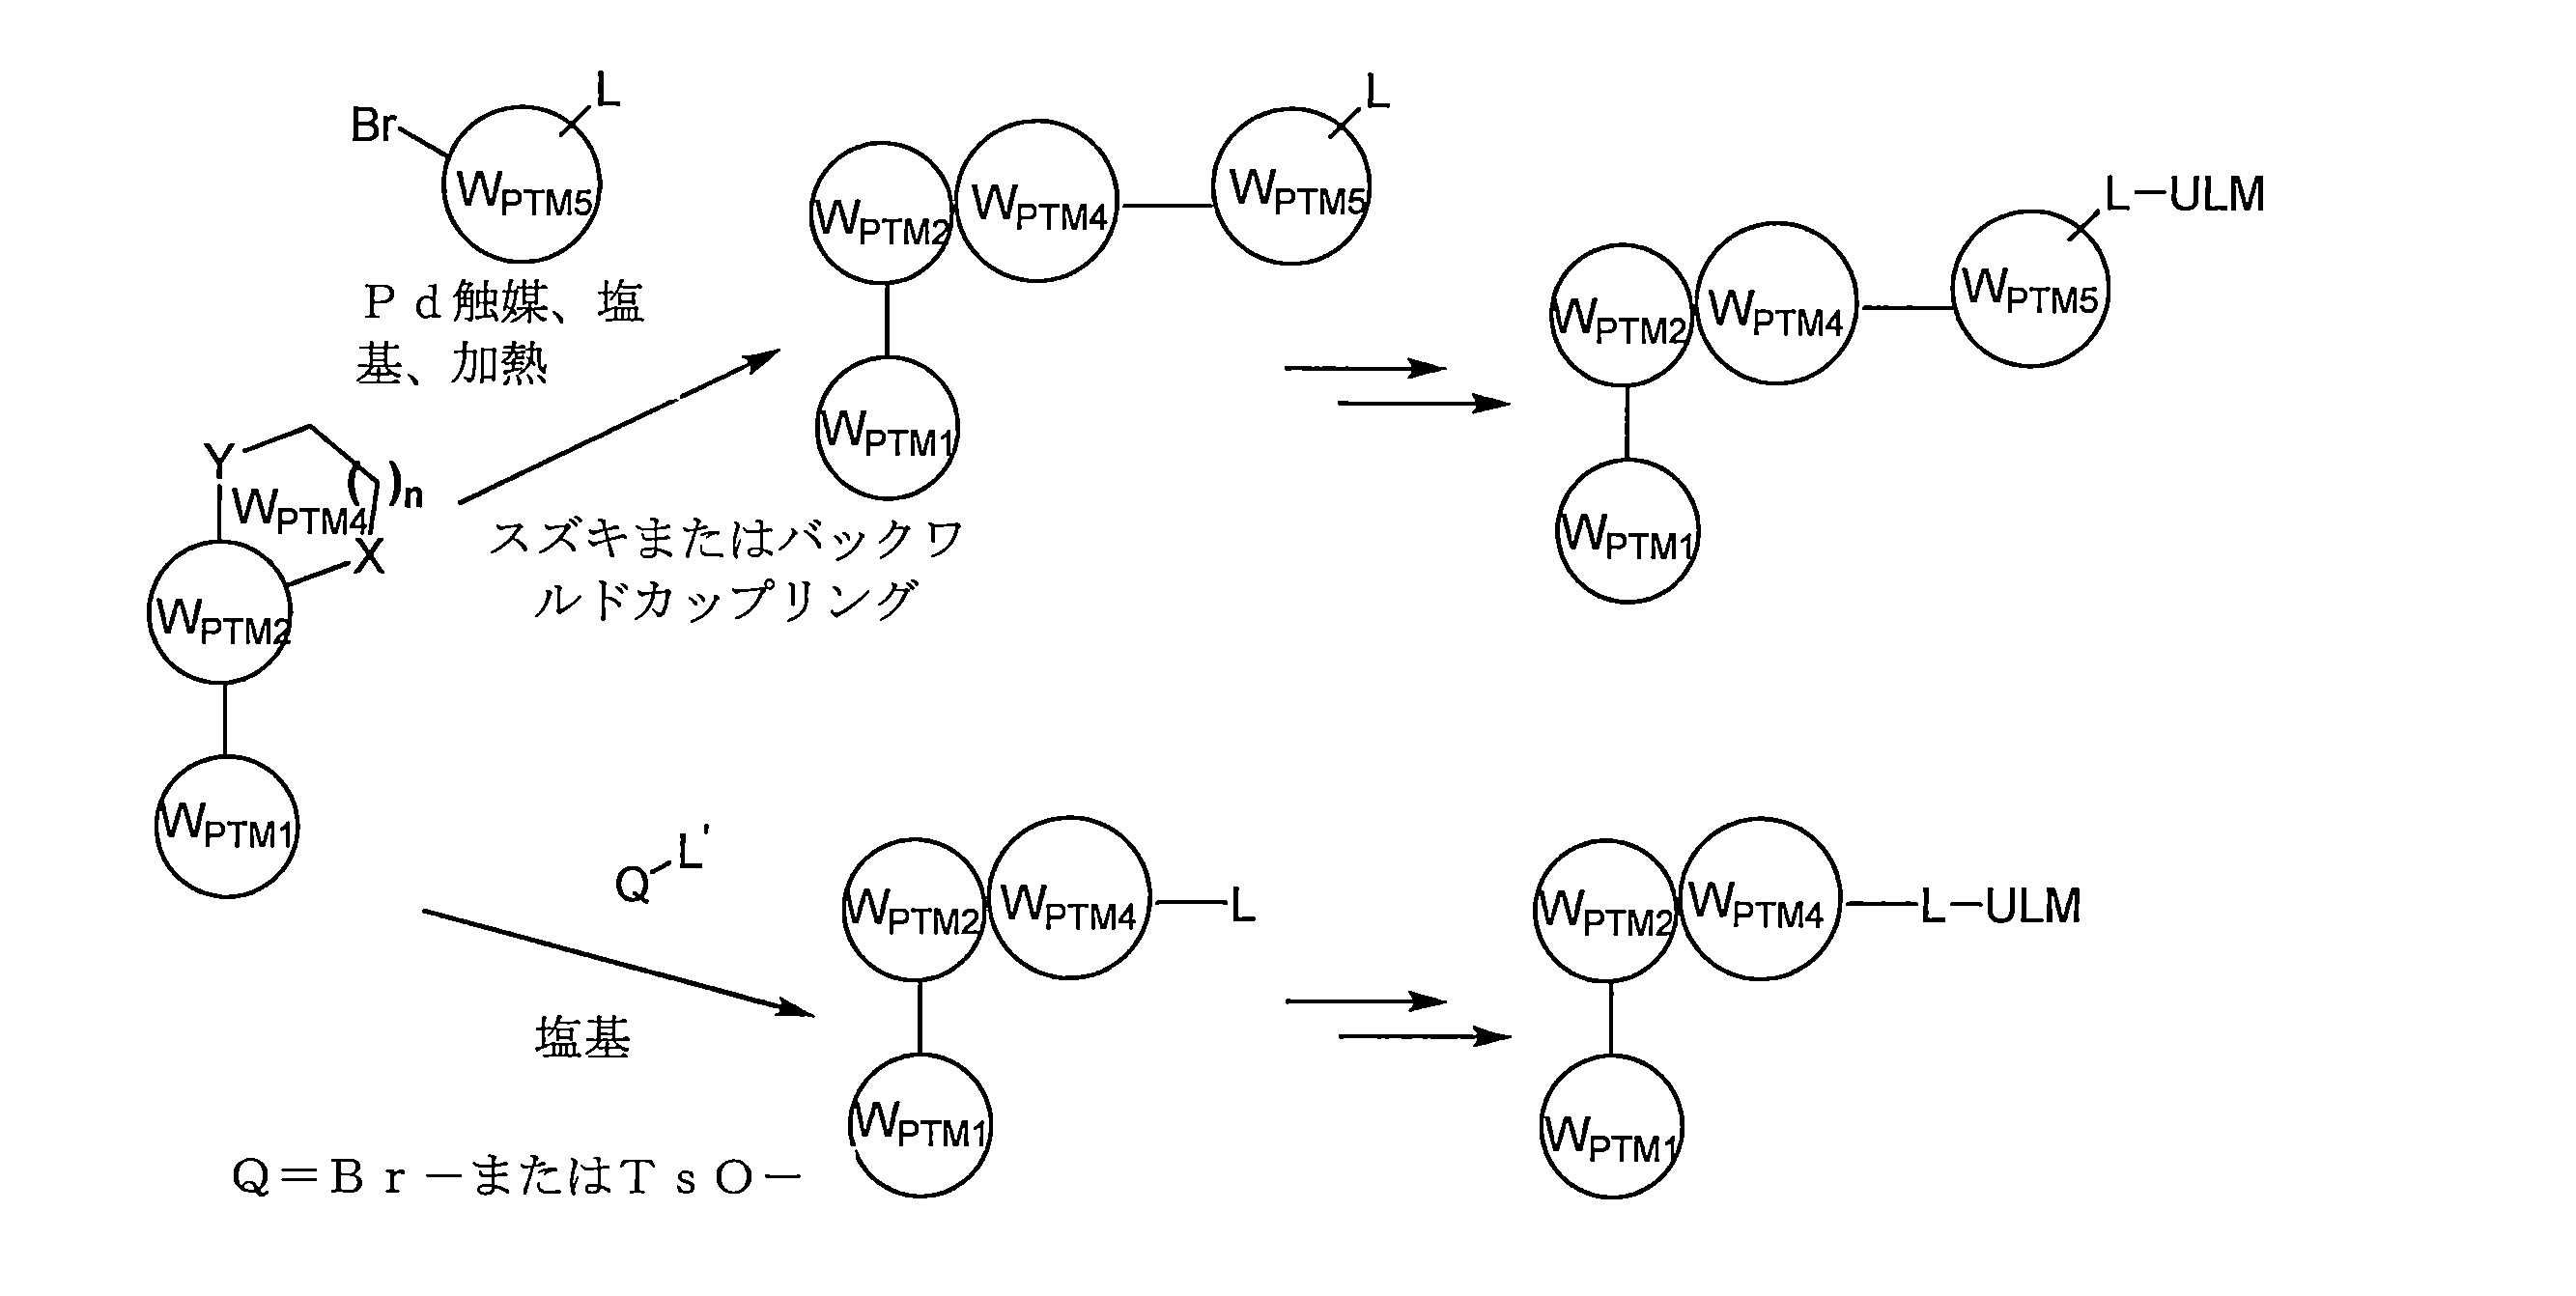
一般式IIIによって表される例示的なPTMは、WPMT5が存在しない場合、式Iの化合物について記載される一般的なアプローチに従って調製することができる。 Exemplary PTMs represented by general formula III can be prepared following the general approach described for compounds of formula I, in the absence of WPMT5.
一般式IVによって表される例示的なPTMは、以下の一般スキームに従って調製することができる。

例示的な化合物11の例示的な合成
工程1
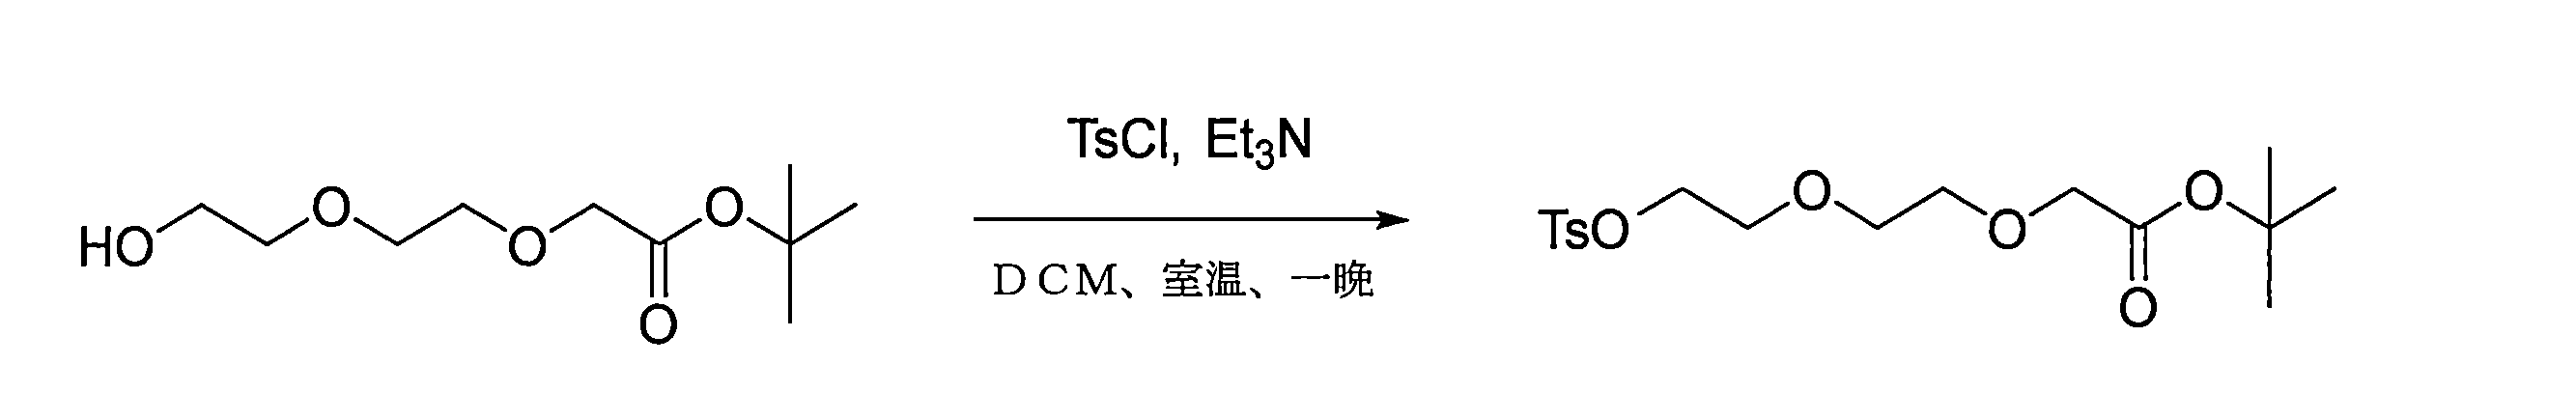
DCM(5mL)中のtert-ブチル2-(2-(2-ヒドロキシエトキシ)エトキシ)酢酸塩(1.5g、6.8mmol)およびTEA(2.07g、20.5mmol)の混合物に、TsCl(1.95g、10.23mmol)を0℃で添加した。得られた混合物を室温に温め、3時間撹拌した。溶液を水(20mL)でクエンチし、DCMで抽出した。有機相をブライン(20mL×2)で洗浄した。有機相をNa2SO4で乾燥させ、濾過して、濃縮した。残渣を、フラッシュクロマトグラフィー(CH2Cl2:MeOH 40:1)によって精製し、tert-ブチル2-(2-(2-(トシルオキシ)エトキシ)エトキシ)酢酸塩(2.14g、mmol、収率84%)を得た。 To a mixture of tert-butyl 2-(2-(2-hydroxyethoxy)ethoxy)acetate (1.5 g, 6.8 mmol) and TEA (2.07 g, 20.5 mmol) in DCM (5 mL) was added TsCl (1.95 g, 10.23 mmol) at 0 °C. The resulting mixture was warmed to room temperature and stirred for 3 h. The solution was quenched with water (20 mL) and extracted with DCM. The organic phase was washed with brine (20 mL x 2). The organic phase was dried over Na2SO4, filtered, and concentrated. The residue was purified by flash chromatography (CH2Cl2:MeOH 40:1) to give tert-butyl 2-(2-(2-(tosyloxy)ethoxy)ethoxy)acetate (2.14 g, mmol, 84% yield).
工程2

乾燥DMF(10mL)中の4-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)-1H-ピラゾール(1.11g、5.722mmol)、tert-ブチル2-(2-(2-(トシルオキシ)エトキシ)エトキシ)酢酸塩(2.14g、5.722mmol)、およびCs2CO3(3.73g、11.444mmol)の混合物を、75℃に3時間加熱した。次いで、反応混合物を室温に冷却し、EtOAc(30mL)で希釈した。有機層を、水(10mL)およびブライン(10mL×2)で洗浄し、乾燥させ(Na2SO4)、濾過して、濃縮した。粗残渣(2.8g)を、さらに精製することなく、次の反応に使用した。 A mixture of 4-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)-1H-pyrazole (1.11 g, 5.722 mmol), tert-butyl 2-(2-(2-(tosyloxy)ethoxy)ethoxy)acetate (2.14 g, 5.722 mmol), and Cs2CO3 (3.73 g, 11.444 mmol) in dry DMF (10 mL) was heated to 75 °C for 3 h. The reaction mixture was then cooled to room temperature and diluted with EtOAc (30 mL). The organic layer was washed with water (10 mL) and brine (10 mL x 2), dried (Na2SO4), filtered, and concentrated. The crude residue (2.8 g) was used in the next reaction without further purification.
工程3

ジオキサン(10mL)および水(1mL)中のtert-ブチル2-(2-(2-(4-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)-1H-ピラゾール-1-イル)エトキシ)エトキシ)酢酸塩(1.5g、3.79mmol)、4-ブロモ-6-クロロピリダジン-3-アミン(1.1g、5.69mmol)、PdCl2(dppf)(555mg、0.758mmol)、tBuPHBF4(441mg、1.52mmol)、およびCs2CO3(3.09g、9.48mmol)の混合物を、N2下で3時間撹拌しながら100℃に加熱した。固形分を濾過し、濾液を濃縮した。残渣を、クロマトグラフィー(CH2Cl2:MeOH 30:1)によって精製して、tert-ブチル2-(2-(2-(4-(3-アミノ-6-クロロピリダジン-4-イル)-1H-ピラゾール-1-イル)エトキシ)エトキシ)酢酸塩(800mg、収率53%)を得た。 A mixture of tert-butyl 2-(2-(2-(4-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)-1H-pyrazol-1-yl)ethoxy)ethoxy)acetate (1.5 g, 3.79 mmol), 4-bromo-6-chloropyridazin-3-amine (1.1 g, 5.69 mmol), PdCl2(dppf) (555 mg, 0.758 mmol), tBuPHBF4 (441 mg, 1.52 mmol), and Cs2CO3 (3.09 g, 9.48 mmol) in dioxane (10 mL) and water (1 mL) was heated to 100 °C with stirring under N2 for 3 h. The solids were filtered and the filtrate was concentrated. The residue was purified by chromatography (CH2Cl2:MeOH 30:1) to give tert-butyl 2-(2-(2-(4-(3-amino-6-chloropyridazin-4-yl)-1H-pyrazol-1-yl)ethoxy)ethoxy)acetate (800 mg, 53% yield).
工程4

ジオキサン(10mL)および水(1mL)中のtert-ブチル2-(2-(2-(4-(3-アミノ-6-クロロピリダジン-4-イル)-1H-ピラゾール-1-イル)エトキシ)エトキシ)酢酸塩(800mg、2.02mmol)、(2-ヒドロキシフェニル)ボロン酸(418mg、3.03mmol)、炭酸セシウム(1.65g、5.05mmol)、PdCl2(dppf)(444mg、0.606mmol)、およびtBu3PHBF4(352mg、1.212mmol)の懸濁液を、窒素下で100℃に3時間加熱した。混合物を室温に冷却し、固形分を濾過した。濾液を濃縮し、クロマトグラフィー(CH2Cl2:MeOH 40:1)によって精製して、tert-ブチル2-(2-(2-(4-(3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル)-1H-ピラゾール-1-イル)エトキシ)エトキシ)酢酸塩(400mg、収率44%)を得た。 A suspension of tert-butyl 2-(2-(2-(4-(3-amino-6-chloropyridazin-4-yl)-1H-pyrazol-1-yl)ethoxy)ethoxy)acetate (800 mg, 2.02 mmol), (2-hydroxyphenyl)boronic acid (418 mg, 3.03 mmol), cesium carbonate (1.65 g, 5.05 mmol), PdCl2(dppf) (444 mg, 0.606 mmol), and tBu3PHBF4 (352 mg, 1.212 mmol) in dioxane (10 mL) and water (1 mL) was heated to 100 °C under nitrogen for 3 h. The mixture was cooled to room temperature and the solids were filtered. The filtrate was concentrated and purified by chromatography (CH2Cl2:MeOH 40:1) to give tert-butyl 2-(2-(2-(4-(3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl)-1H-pyrazol-1-yl)ethoxy)ethoxy)acetate (400 mg, 44% yield).
工程5

THF/H2O(5mL、2:1)中のtert-ブチル2-(2-(2-(4-(3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル)-1H-ピラゾール-1-イル)エトキシ)エトキシ)酢酸塩(400mg、0.88mmol)の溶液に、LiOH(111mg、2.64mmol)を0℃で添加した。混合物を0℃で2時間撹拌した。反応溶液を、1MのHClでクエンチした。溶液を乾燥(Na2SO4)させ、濾過し、減圧下で濃縮して、粗2-(2-(2-(4-(3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル)-1H-ピラゾール-1-イル)エトキシ)エトキシ)酢酸(600mg)を得、これを、さらに精製することなく次の工程で使用した。 To a solution of tert-butyl 2-(2-(2-(4-(3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl)-1H-pyrazol-1-yl)ethoxy)ethoxy)acetate (400 mg, 0.88 mmol) in THF/H2O (5 mL, 2:1) was added LiOH (111 mg, 2.64 mmol) at 0°C. The mixture was stirred at 0°C for 2 h. The reaction solution was quenched with 1M HCl. The solution was dried (Na2SO4), filtered, and concentrated under reduced pressure to give crude 2-(2-(2-(4-(3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl)-1H-pyrazol-1-yl)ethoxy)ethoxy)acetic acid (600 mg), which was used in the next step without further purification.
工程6

DMF(5mL)中の2-(2-(2-(4-(3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル)-1H-ピラゾール-1-イル)エトキシ)エトキシ)酢酸(200mg粗製)、(2S,4R)-1-((S)-2-アミノ-3,3-ジメチルブタノイル)-4-ヒドロキシ-N-(4-(4-メチルチアゾール-5-イル)ベンジル)ピロリジン-2-カルボキサミド(430mg、1mmol)、およびDIPEA(516mg、4mmol)の溶液に、HATU(570mg、1.5mmol)を0℃で添加した。反応混合物を室温で30分間撹拌した。混合物を酢酸エチル(50mL)で抽出した。合わせた有機層をブライン(8mL×2)で洗浄し、乾燥(Na2SO4)させて、濾過した。有機層を減圧下で濃縮した。残渣を、分取TLC(DCM中の6%MeOH)によって精製して、(2S,4R)-1-((S)-2-(2-(2-(2-(4-(3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル)-1H-ピラゾール-1-イル)エトキシ)エトキシ)アセトアミド)-3,3-ジメチルブタノイル)-4-ヒドロキシ-N-(4-(4-メチルチアゾール-5-イル)ベンジル)ピロリジン-2-カルボキサミド(22mg)を得た。 To a solution of 2-(2-(2-(4-(3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl)-1H-pyrazol-1-yl)ethoxy)ethoxy)acetic acid (200 mg crude), (2S,4R)-1-((S)-2-amino-3,3-dimethylbutanoyl)-4-hydroxy-N-(4-(4-methylthiazol-5-yl)benzyl)pyrrolidine-2-carboxamide (430 mg, 1 mmol), and DIPEA (516 mg, 4 mmol) in DMF (5 mL) was added HATU (570 mg, 1.5 mmol) at 0°C. The reaction mixture was stirred at room temperature for 30 min. The mixture was extracted with ethyl acetate (50 mL). The combined organic layers were washed with brine (8 mL x 2), dried (Na2SO4), and filtered. The organic layers were concentrated under reduced pressure. The residue was purified by preparative TLC (6% MeOH in DCM) to give (2S,4R)-1-((S)-2-(2-(2-(2-(4-(3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl)-1H-pyrazol-1-yl)ethoxy)ethoxy)acetamido)-3,3-dimethylbutanoyl)-4-hydroxy-N-(4-(4-methylthiazol-5-yl)benzyl)pyrrolidine-2-carboxamide (22 mg).
1HNMR(400MHz、MeOD):δ8.83(s、1H)、8.32(s、1H)、8.04(d、J=9.6Hz、2H)、7.78(d、J=7.6Hz、1H)、7.65(d、J=10.0Hz、1H)、7.39(d、J=8.0Hz、2H)、7.32~7.34(m、2H)、7.28(m、1H)、6.95(d、J=7.6Hz 2H)、4.69(d、J=9.6Hz 1H)、4.44~4.57(m、6H)、4.25~4.35(m、1H)、3.60~3.99(m、11H)、2.46(s、3H)、2.23(m、1H)、2.08(m、1H)、1.01(s、9H)。 1H NMR (400MHz, MeOD): δ 8.83 (s, 1H), 8.32 (s, 1H), 8.04 (d, J = 9.6Hz, 2H), 7.78 (d, J = 7.6Hz, 1H), 7.65 (d, J = 10.0Hz, 1H), 7.39 (d, J = 8.0Hz, 2H), 7.32-7.34 (m, 2H), 7.28 (m, 1H), 6.95 (d, J = 7.6Hz 2H), 4.69 (d, J = 9.6Hz 1H), 4.44-4.57 (m, 6H), 4.25-4.35 (m, 1H), 3.60-3.99 (m, 11H), 2.46 (s, 3H), 2.23 (m, 1H), 2.08 (m, 1H), 1.01 (s, 9H).
例示的な化合物5を、例示的な化合物11について上に記載されるものと類似の手順、ならびに当業者に既知であり、かつ理解される手順を使用して、以下のスキームに従って調製した。

例示的な化合物11および例示的な化合物5について記載される手順を使用して、例示的な化合物1、例示的な化合物2、例示的な化合物3、例示的な化合物4を調製した。 Using the procedures described for Exemplary Compound 11 and Exemplary Compound 5, Exemplary Compound 1, Exemplary Compound 2, Exemplary Compound 3, Exemplary Compound 4 were prepared.
例示的な化合物9の例示的な合成
工程1

無水DMF(30mL)中の3,6,9,12-テトラオキサテトラデカン-1,14-ジオール(13.5g、56.8mmol)の溶液に、60%NaH(1.25g、31.2mmol)を0℃で添加した。反応混合物を室温で0.5時間撹拌した。次いで、4-ブロモ-2-フルオロピリジン(5g、28.4mmol)を、混合物に滴下で添加し、混合物を75℃に2時間加熱した。反応混合物を水(10mL)でクエンチし、EA(200mL)で抽出した。有機相をブライン(10mL)で洗浄し、乾燥(Na2SO4)させて、減圧下で濃縮した。残渣を、シリカゲルカラムによって精製して、14-((4-ブロモピリジン-2-イル)オキシ)-3,6,9,12-テトラオキサテトラデカン-1-オール(9.0g、22.9mmol、収率81%)を得た。 To a solution of 3,6,9,12-tetraoxatetradecane-1,14-diol (13.5 g, 56.8 mmol) in anhydrous DMF (30 mL) was added 60% NaH (1.25 g, 31.2 mmol) at 0 °C. The reaction mixture was stirred at room temperature for 0.5 h. Then, 4-bromo-2-fluoropyridine (5 g, 28.4 mmol) was added dropwise to the mixture and the mixture was heated to 75 °C for 2 h. The reaction mixture was quenched with water (10 mL) and extracted with EA (200 mL). The organic phase was washed with brine (10 mL), dried (Na2SO4) and concentrated under reduced pressure. The residue was purified by silica gel column to give 14-((4-bromopyridin-2-yl)oxy)-3,6,9,12-tetraoxatetradecane-1-ol (9.0 g, 22.9 mmol, 81% yield).
工程2

無水THF(50mL)中の14-((4-ブロモピリジン-2-イル)オキシ)-3,6,9,12-テトラオキサテトラデカン-1-オール(5.0g、12.7mmol)の溶液に、60%NaH(660mg、16.5mmol)を0℃で添加した。反応混合物を室温で40分間撹拌した。次いで、tert-ブチル2-ブロモ酢酸塩(4.9g、25.4mmol)を、混合物に滴下で添加し、室温で一晩撹拌した。反応混合物を2N NH4Cl(10mL)でクエンチし、EA(200mL)で抽出した。次いで、有機相をブライン(10mL)で洗浄し、乾燥(Na2SO4)させて、減圧下で濃縮した。残渣を、シリカゲルカラムによって精製して、tert-ブチル17-((4-ブロモピリジン-2-イル)オキシ)-3,6,9,12,15-ペンタオキサヘプタデカノエート(2.6g、5.13mmol、収率40%)を得た。 To a solution of 14-((4-bromopyridin-2-yl)oxy)-3,6,9,12-tetraoxatetradecan-1-ol (5.0 g, 12.7 mmol) in anhydrous THF (50 mL) was added 60% NaH (660 mg, 16.5 mmol) at 0 °C. The reaction mixture was stirred at room temperature for 40 min. Then, tert-butyl 2-bromoacetate (4.9 g, 25.4 mmol) was added dropwise to the mixture and stirred at room temperature overnight. The reaction mixture was quenched with 2N NH4Cl (10 mL) and extracted with EA (200 mL). The organic phase was then washed with brine (10 mL), dried (Na2SO4) and concentrated under reduced pressure. The residue was purified by silica gel column to obtain tert-butyl 17-((4-bromopyridin-2-yl)oxy)-3,6,9,12,15-pentaoxaheptadecanoate (2.6 g, 5.13 mmol, 40% yield).
工程3
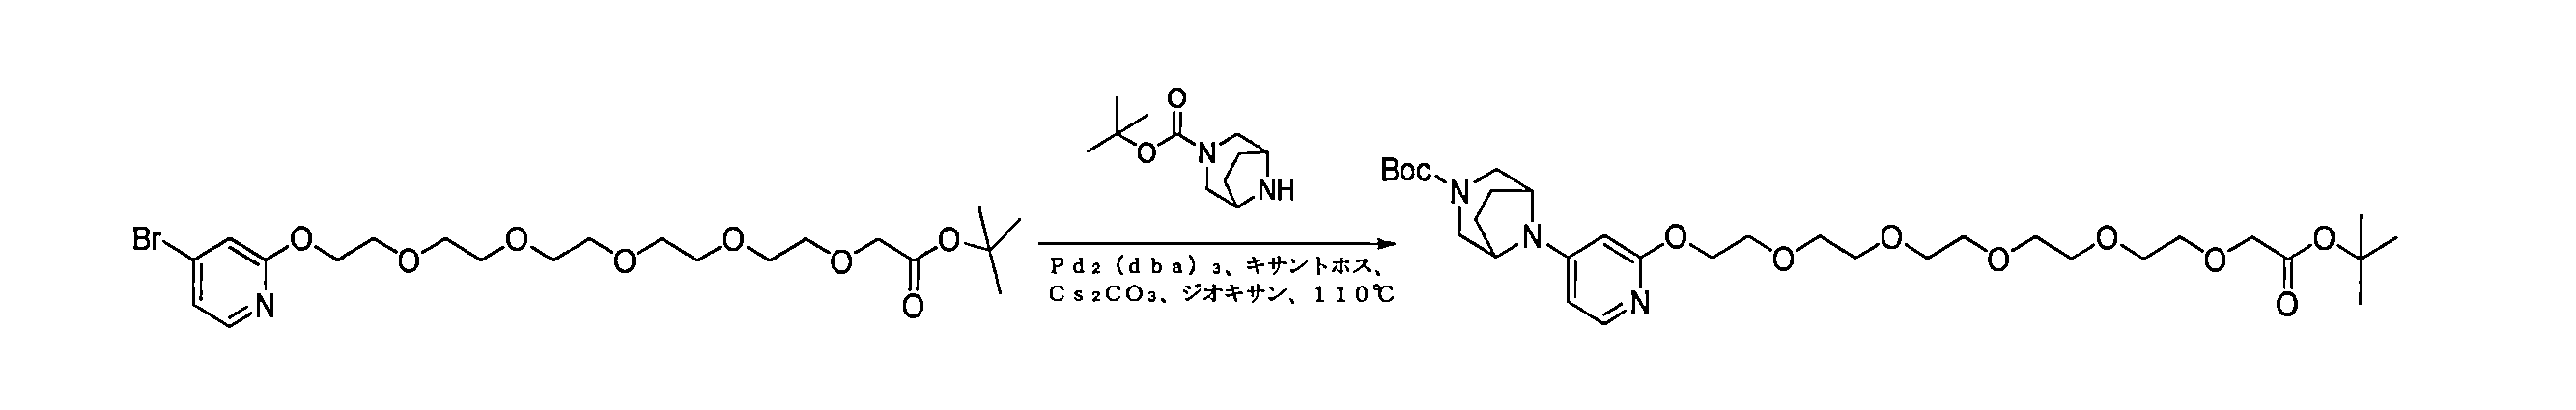
密封チューブ中のジオキサン(5mL)中のtert-ブチル17-((4-ブロモピリジン-2-イル)オキシ)-3,6,9,12,15-ペンタオキサヘプタデカノエート(250mg、1.18mmol)、tert-ブチル3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレート(718mg、1.4mmol)、炭酸セシウム(769mg、2.36mmol)、Pd2(dba)3(110mg、0.12mmol)、およびキサントホス(138mg、0.24mmol)の混合物を、窒素下で110℃に一晩加熱した。混合物をEA(100mL)で抽出した。次いで、有機相を、水(10mL)、ブライン(10mL)で洗浄し、乾燥(Na2SO4)させて、減圧下で濃縮した。残渣を、シリカゲルカラムによって精製して、tert-ブチル8-(2-((19,19-ジメチル-17-オキソ-3,6,9,12,15,18-ヘキサオキサイコシル)オキシ)ピリジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレート(550mg、0.86mmol、収率73%)を得た。 A mixture of tert-butyl 17-((4-bromopyridin-2-yl)oxy)-3,6,9,12,15-pentaoxaheptadecanoate (250 mg, 1.18 mmol), tert-butyl 3,8-diazabicyclo[3.2.1]octane-3-carboxylate (718 mg, 1.4 mmol), cesium carbonate (769 mg, 2.36 mmol), Pd2(dba)3 (110 mg, 0.12 mmol), and Xantphos (138 mg, 0.24 mmol) in dioxane (5 mL) in a sealed tube was heated to 110 °C overnight under nitrogen. The mixture was extracted with EA (100 mL). The organic phase was then washed with water (10 mL), brine (10 mL), dried (Na2SO4), and concentrated under reduced pressure. The residue was purified by silica gel column to obtain tert-butyl 8-(2-((19,19-dimethyl-17-oxo-3,6,9,12,15,18-hexaoxaicosyl)oxy)pyridin-4-yl)-3,8-diazabicyclo[3.2.1]octane-3-carboxylate (550 mg, 0.86 mmol, yield 73%).
工程4

MeOH(15mL)中のtert-ブチル8-(2-((19,19-ジメチル-17-オキソ-3,6,9,12,15,18-ヘキサオキサイコシル)オキシ)ピリジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレート(550mg、0.86mmol)の溶液に、ジオキサン中のHCl(ジオキサン中6N)(5mL、30mmol)を室温で添加した。反応混合物を、室温で1時間撹拌した。混合物を減圧下で濃縮して、粗メチル17-((4-(3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル)オキシ)-3,6,9,12,15-ペンタオキサヘプタデカノエート(0.55g)を得た。 To a solution of tert-butyl 8-(2-((19,19-dimethyl-17-oxo-3,6,9,12,15,18-hexaoxaicosyl)oxy)pyridin-4-yl)-3,8-diazabicyclo[3.2.1]octane-3-carboxylate (550 mg, 0.86 mmol) in MeOH (15 mL) was added HCl in dioxane (6N in dioxane) (5 mL, 30 mmol) at room temperature. The reaction mixture was stirred at room temperature for 1 hour. The mixture was concentrated under reduced pressure to give crude methyl 17-((4-(3,8-diazabicyclo[3.2.1]octane-8-yl)pyridin-2-yl)oxy)-3,6,9,12,15-pentaoxaheptadecanoate (0.55 g).
工程5
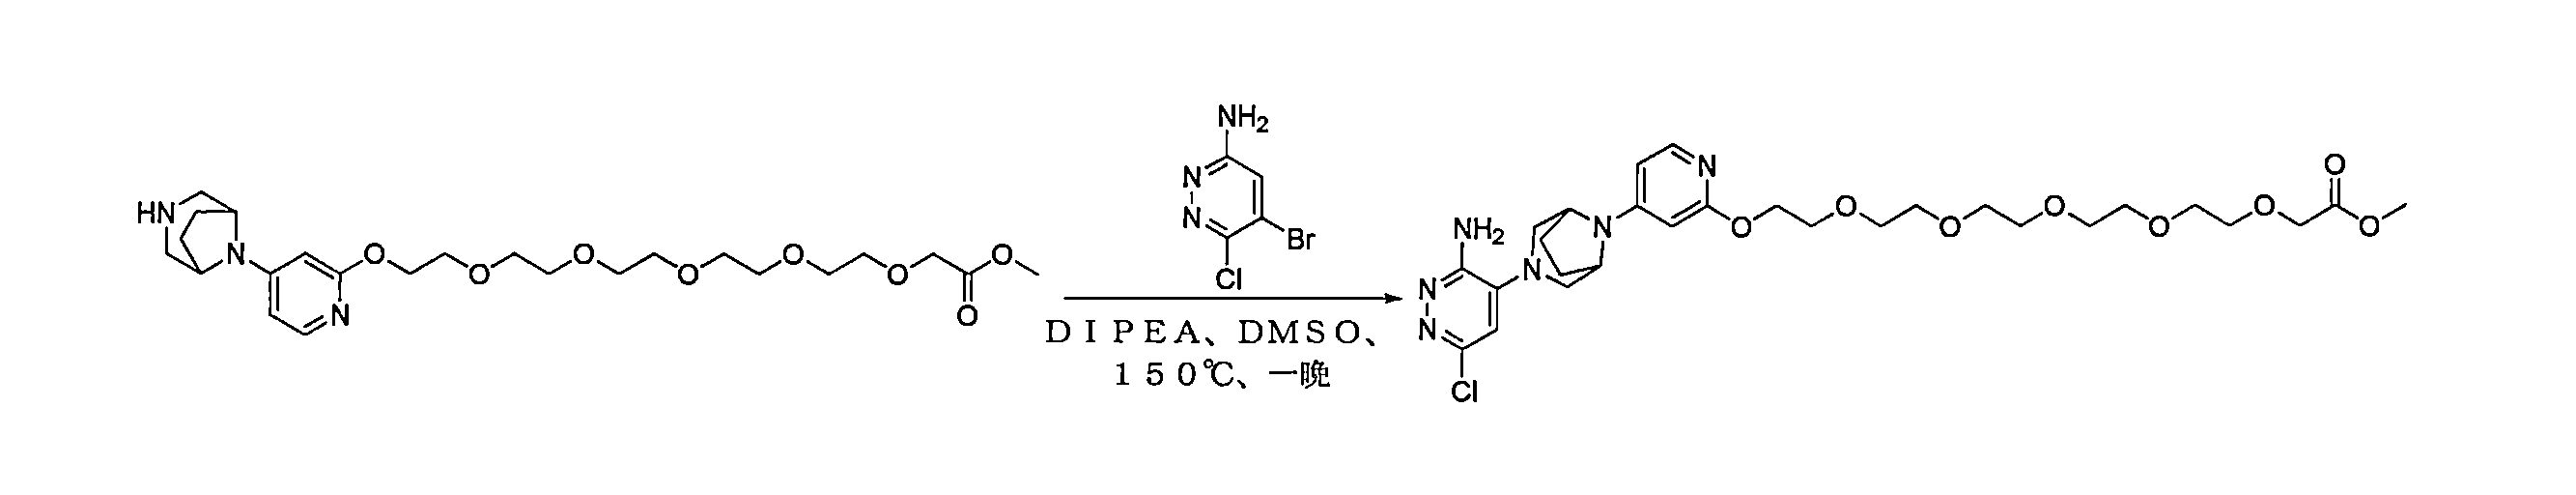
DMSO(5mL)中の粗メチル17-((4-(3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル)オキシ)-3,6,9,12,15-ペンタオキサヘプタデカノエート(550mg、粗製)の溶液に、5-ブロモ-6-クロロピリダジン-3-アミン(526mg、2.71mmol)およびDIPEA(1.87g、14.5mmol)を添加した。溶液を、150℃で一晩撹拌した。混合物をEA(60mL)で抽出した。有機相を水(8mL)およびブライン(8mL)で洗浄した。有機層を乾燥させ(Na2SO4)、減圧下で濾過および濃縮した。残渣を、シリカゲルカラムクロマトグラフィーによって精製して、メチル17-((4-((1R,5S)-3-(3-アミノ-6-クロロピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル)オキシ)-3,6,9,12,15-ペンタオキサヘプタデカノエート(400mg)を得た。 To a solution of crude methyl 17-((4-(3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl)oxy)-3,6,9,12,15-pentaoxaheptadecanoate (550 mg, crude) in DMSO (5 mL) was added 5-bromo-6-chloropyridazin-3-amine (526 mg, 2.71 mmol) and DIPEA (1.87 g, 14.5 mmol). The solution was stirred at 150 °C overnight. The mixture was extracted with EA (60 mL). The organic phase was washed with water (8 mL) and brine (8 mL). The organic layer was dried (Na2SO4), filtered and concentrated under reduced pressure. The residue was purified by silica gel column chromatography to obtain methyl 17-((4-((1R,5S)-3-(3-amino-6-chloropyridazin-4-yl)-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl)oxy)-3,6,9,12,15-pentaoxaheptadecanoate (400 mg).
工程6

ジオキサン(12mL)および水(1.2mL)中のメチル17-((4-((1R,5S)-3-(3-アミノ-6-クロロピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル)オキシ)-3,6,9,12,15-ペンタオキサヘプタデカノエート(325mg、0.52mmol)および(2-ヒドロキシフェニル)ボロン酸(93mg、0.68mmol)の溶液に、炭酸セシウム(542mg、1.66mmol)、PdCl2(dppf)(73.2mg、0.1mmol)、およびt-Bu3PHBF4(58mg、0.2mmol)を添加した。溶液を、窒素雰囲気下で、100℃で4時間撹拌した。溶液のpHを、1N HClで5に調整した。混合物を濾過し、濾液を減圧下で濃縮して、粗17-((4-((1R,5S)-3-(3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル)オキシ)-3,6,9,12,15-ペンタオキサヘプタデカン酸(300mg)を得、これを、さらに精製することなく次の工程で使用した。 To a solution of methyl 17-((4-((1R,5S)-3-(3-amino-6-chloropyridazin-4-yl)-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl)oxy)-3,6,9,12,15-pentaoxaheptadecanoate (325 mg, 0.52 mmol) and (2-hydroxyphenyl)boronic acid (93 mg, 0.68 mmol) in dioxane (12 mL) and water (1.2 mL) was added cesium carbonate (542 mg, 1.66 mmol), PdCl2(dppf) (73.2 mg, 0.1 mmol), and t-Bu3PHBF4 (58 mg, 0.2 mmol). The solution was stirred at 100°C for 4 h under nitrogen atmosphere. The pH of the solution was adjusted to 5 with 1N HCl. The mixture was filtered and the filtrate was concentrated under reduced pressure to give crude 17-((4-((1R,5S)-3-(3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl)-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl)oxy)-3,6,9,12,15-pentaoxaheptadecanoic acid (300 mg), which was used in the next step without further purification.
工程7

DMF(5mL)中の粗17-((4-((1R,5S)-3-(3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル)オキシ)-3,6,9,12,15-ペンタオキサヘプタデカン酸(80mg、0.12mmol)および(2S,4R)-1-((S)-2-アミノ-3,3-ジメチルブタニル)-4-ヒドロキシ-N-(4-(4-メチルチアゾール-5-イル)ベンジル)ピロリジン-2-カルボキサミド(108mg、0.24mmol)の溶液に、DIPEA(124mg、0.96mmol)およびHATU(92mg、0.24mmol)を0℃で添加した。反応混合物を室温で30分間撹拌した。混合物をEA(50mL)で抽出した。有機相を水(8mL)およびブライン(8mL)で洗浄した。有機層を乾燥させ(Na2SO4)、減圧下で濾過および濃縮した。残渣を、分取TLC(DCM中6%MeOH)によって精製して、(2S,4R)-1-((S)-20-((4-((1R,5S)-3-(3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル)オキシ)-2-(tert-ブチル)-4-オキソ-6,9,12,15,18-ペンタオキサ-3-アザイコサノイル)-4-ヒドロキシ-N-(4-(4-メチルチアゾール-5-イル)ベンジル)ピロリジン-2-カルボキサミド(20mg、0.018mmol、収率15.0%)を得た。 To a solution of crude 17-((4-((1R,5S)-3-(3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl)-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl)oxy)-3,6,9,12,15-pentaoxaheptadecanoic acid (80 mg, 0.12 mmol) and (2S,4R)-1-((S)-2-amino-3,3-dimethylbutanyl)-4-hydroxy-N-(4-(4-methylthiazol-5-yl)benzyl)pyrrolidine-2-carboxamide (108 mg, 0.24 mmol) in DMF (5 mL) was added DIPEA (124 mg, 0.96 mmol) and HATU (92 mg, 0.24 mmol) at 0 °C. The reaction mixture was stirred at room temperature for 30 min. The mixture was extracted with EA (50 mL). The organic phase was washed with water (8 mL) and brine (8 mL). The organic layer was dried (Na2SO4), filtered and concentrated under reduced pressure. The residue was purified by preparative TLC (6% MeOH in DCM) to give (2S,4R)-1-((S)-20-((4-((1R,5S)-3-(3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl)-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl)oxy)-2-(tert-butyl)-4-oxo-6,9,12,15,18-pentaoxa-3-azaicosanoyl)-4-hydroxy-N-(4-(4-methylthiazol-5-yl)benzyl)pyrrolidine-2-carboxamide (20 mg, 0.018 mmol, 15.0% yield).
1H NMR(400MHz、MeOD):δ8.74(s、1H)、7.68~7.66(m、2H)、7.34~7.28(m、5H)、7.18~7.10(m、1H)、6.80~6.78(m、2H)、6.43~6.40(m、1H)、6.10(s、1H)、4.60~4.20(m、9H)、3.92~3.89(m、2H)、3.80~3.64(m、4H)、3.71~3.45(m、16H)、3.10~3.00(m、1H)、3.00~2.90(m、2H)、2.37~2.34(m、1H)、2.35(s、3H)、2.18~1.99(m、6H)、0.92(s、9H)。 1H NMR (400MHz, MeOD): δ 8.74 (s, 1H), 7.68-7.66 (m, 2H), 7.34-7.28 (m, 5H), 7.18-7.10 (m, 1H), 6.80-6.78 (m, 2H), 6.43-6.40 (m, 1H), 6.10 (s, 1H), 4.60-4.20 (m , 9H), 3.92-3.89 (m, 2H), 3.80-3.64 (m, 4H), 3.71-3.45 (m, 16H), 3.10-3.00 (m, 1H ), 3.00-2.90 (m, 2H), 2.37-2.34 (m, 1H), 2.35 (s, 3H), 2.18-1.99 (m, 6H), 0.92 (s, 9H).
類似の手順を使用して、例示的な化合物7、8、10、および35を調製した。 Using similar procedures, exemplary compounds 7, 8, 10, and 35 were prepared.
例示的な化合物6の例示的な合成
工程1

DCM(50mL)およびH2O(25mL)中の2-(2-(2-(2-((4-ブロモピリジン-2-イル)オキシ)エトキシ)エトキシ)エトキシ)エタン-1-オール(3.49g、10mmol)の溶液[実施例A2979について記載されるものと類似の手順を使用して調製]に、PhI(OAc)2(9.66g、30mmol)およびTEMPO(312mg、2mmol)を添加した。反応混合物を、室温で2時間撹拌した。混合物をEA(100mL)で抽出した。有機相を水(10mL)およびブライン(10mL)で洗浄した。有機層を乾燥させ(Na2SO4)、減圧下で濾過および濃縮した。残渣を、シリカゲルカラムクロマトグラフィーによって精製して、2-(2-(2-(2-((4-ブロモピリジン-2-イル)オキシ)エトキシ)エトキシ)エトキシ)酢酸(3.5g、9.64mmol、収率96%)を得た。 To a solution of 2-(2-(2-(2-((4-bromopyridin-2-yl)oxy)ethoxy)ethoxy)ethoxy)ethan-1-ol (3.49 g, 10 mmol) [prepared using a procedure similar to that described for Example A2979] in DCM (50 mL) and H2O (25 mL) was added PhI(OAc)2 (9.66 g, 30 mmol) and TEMPO (312 mg, 2 mmol). The reaction mixture was stirred at room temperature for 2 h. The mixture was extracted with EA (100 mL). The organic phase was washed with water (10 mL) and brine (10 mL). The organic layer was dried (Na2SO4), filtered and concentrated under reduced pressure. The residue was purified by silica gel column chromatography to obtain 2-(2-(2-(2-((4-bromopyridin-2-yl)oxy)ethoxy)ethoxy)ethoxy)acetic acid (3.5 g, 9.64 mmol, yield 96%).
工程2

MeOH(30mL)中の2-(2-(2-(2-((4-ブロモピリジン-2-イル)オキシ)エトキシ)エトキシ)エトキシ)酢酸(3.0g、8.26mmol)の溶液に、SOCl2(4.0g、33.9mmol)を0℃において滴下で添加した。反応混合物を室温で4時間撹拌した。溶液のpHを、飽和NaHCO3で約8に調整した。混合物をDCM(100mL)で抽出した。有機相を水(10mL)およびブライン(10mL)で洗浄した。有機層を乾燥(Na2SO4)し、濾過し、減圧下で濃縮して、メチル2-(2-(2-(2-((4-ブロモピリジン-2-イル)オキシ)エトキシ)エトキシ)エトキシ)酢酸塩(2.9g、7.69mmol、収率93%)を得た。 To a solution of 2-(2-(2-(2-((4-bromopyridin-2-yl)oxy)ethoxy)ethoxy)ethoxy)acetic acid (3.0 g, 8.26 mmol) in MeOH (30 mL) was added SOCl2 (4.0 g, 33.9 mmol) dropwise at 0 °C. The reaction mixture was stirred at room temperature for 4 h. The pH of the solution was adjusted to about 8 with saturated NaHCO3. The mixture was extracted with DCM (100 mL). The organic phase was washed with water (10 mL) and brine (10 mL). The organic layer was dried (Na2SO4), filtered and concentrated under reduced pressure to give methyl 2-(2-(2-(2-((4-bromopyridin-2-yl)oxy)ethoxy)ethoxy)ethoxy)acetate (2.9 g, 7.69 mmol, 93% yield).
例示的な化合物9について記載されるものと類似の手順を使用して、メチル2-(2-(2-(2-((4-ブロモピリジン-2-イル)オキシ)エトキシ)エトキシ)エトキシ)酢酸塩を、最終化合物(2S,4R)-1-((2S)-14-((4-(3-(3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル)オキシ)-2-(tert-ブチル)-4-オキソ-6,9,12-トリオキサ-3-アザテトラデカノイル)-4-ヒドロキシ-N-(4-(4-メチルチアゾール-5-イル)ベンジル)ピロリジン-2-カルボキサミド(例示的な化合物6)に変換した。 Using a procedure similar to that described for exemplary compound 9, methyl 2-(2-(2-(2-((4-bromopyridin-2-yl)oxy)ethoxy)ethoxy)ethoxy)acetate was converted to the final compound (2S,4R)-1-((2S)-14-((4-(3-(3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl)-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl)oxy)-2-(tert-butyl)-4-oxo-6,9,12-trioxa-3-azatetradecanoyl)-4-hydroxy-N-(4-(4-methylthiazol-5-yl)benzyl)pyrrolidine-2-carboxamide (exemplary compound 6).
1HNMR(400MHz、MeOD):δ8.82(s、1H)、7.76~7.74(m、2H)、7.42~7.34(m、5H)、7.22~7.18(m、1H)、6.89~6.86(m、2H)、6.53~6.52(m、1H)、6.16(s、1H)、4.66~4.28(m、9H)、4.01~4.00(m、2H)、3.82~3.60(m、12H)、3.31~3.29(m、1H)、3.10~3.08(m、2H)、2.51~2.50(m、1H)、2.49(s、3H)、2.29~2.05(m、6H)、2.42~2.29(m、2H)、1.01(s、9H)。 1H NMR (400 MHz, MeOD): δ 8.82 (s, 1H), 7.76-7.74 (m, 2H), 7.42-7.34 (m, 5H), 7.22-7.18 ( m, 1H), 6.89-6.86 (m, 2H), 6.53-6.52 (m, 1H), 6.16 (s, 1H), 4.66-4.28 (m, 9H) , 4.01-4.00 (m, 2H), 3.82-3.60 (m, 12H), 3.31-3.29 (m, 1H), 3.10-3.08 (m, 2H), 2.51-2.50 (m, 1H), 2.49 (s, 3H), 2.29-2.05 (m, 6H), 2.42-2.29 (m, 2H) , 1.01 (s, 9H).
例示的な化合物20の例示的な合成
工程1

DCM(50mL)中の3,6,9,12,15-ペンタオキサヘプタデカン-1,17-ジオール(4g、14.2mmol)およびEt3N(8.6g、85.2mmol)の溶液に、TsCl(8.1g、42.6mmol)を添加した。反応混合物を、室温で1時間撹拌した。混合物をEtOAc(100mL)と水(10mL)とに分割した。有機相をブライン(10mL)で洗浄した。合わせた有機層を乾燥させ(Na2SO4)、濾過し、減圧下で濃縮した。残渣を、シリカゲルカラムクロマトグラフィーによって精製して、3,6,9,12,15-ペンタオキサヘプタデカン-1,17-ジイルビス(4-メチルベンゼンスルホネート)(6.0g、10.2mmol、収率72%)を得た。 To a solution of 3,6,9,12,15-pentaoxaheptadecane-1,17-diol (4 g, 14.2 mmol) and Et3N (8.6 g, 85.2 mmol) in DCM (50 mL) was added TsCl (8.1 g, 42.6 mmol). The reaction mixture was stirred at room temperature for 1 h. The mixture was partitioned between EtOAc (100 mL) and water (10 mL). The organic phase was washed with brine (10 mL). The combined organic layers were dried (Na2SO4), filtered, and concentrated under reduced pressure. The residue was purified by silica gel column chromatography to give 3,6,9,12,15-pentaoxaheptadecane-1,17-diylbis(4-methylbenzenesulfonate) (6.0 g, 10.2 mmol, 72% yield).
工程2

無水DMF(20mL)中の3,6,9,12,15-ペンタオキサヘプタデカン-1,17-ジイルビス(4-メチルベンゼンスルホネート)(3.5g、5.93mmol)の溶液に、4-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)-1H-ピラゾール(1.15g、5.93mmol)およびCs2CO3(3.87g、11.86mmol)を添加した。反応混合物を、75℃で0.5時間撹拌した。混合物を室温に冷却し、EtOAc(200mL)と水(20mL)とに分割した。有機相をブライン(20mL)で洗浄した。合わせた有機層を乾燥させ(Na2SO4)、濾過し、減圧下で濃縮した。残渣を、シリカゲルカラムクロマトグラフィーによって精製して、17-(4-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)-1H-ピラゾール-1-イル)-3,6,9,12,15-ペンタオキサヘプタデシル4-メチルベンゼンスルホネート(0.6g、0.98mmol、収率16.5%)を得た。 To a solution of 3,6,9,12,15-pentaoxaheptadecane-1,17-diylbis(4-methylbenzenesulfonate) (3.5 g, 5.93 mmol) in anhydrous DMF (20 mL) was added 4-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)-1H-pyrazole (1.15 g, 5.93 mmol) and Cs2CO3 (3.87 g, 11.86 mmol). The reaction mixture was stirred at 75 °C for 0.5 h. The mixture was cooled to room temperature and partitioned between EtOAc (200 mL) and water (20 mL). The organic phase was washed with brine (20 mL). The combined organic layers were dried (Na2SO4), filtered, and concentrated under reduced pressure. The residue was purified by silica gel column chromatography to obtain 17-(4-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)-1H-pyrazol-1-yl)-3,6,9,12,15-pentaoxaheptadecyl 4-methylbenzenesulfonate (0.6 g, 0.98 mmol, yield 16.5%).
工程3

DMF(5mL)中の17-(4-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)-1H-ピラゾール-1-イル)-3,6,9,12,15-ペンタオキサヘプタデシル4-メチルベンゼンスルホネート(0.3g、0.49mmol)および(2S,4R)-4-ヒドロキシ-N-(2-ヒドロキシ-4-(4-メチルチアゾール-5-イル)ベンジル)-1-((S)-3-メチル-2-(1-オキソイソインドリン-2-イル)ブタノイル)ピロリジン-2-カルボキサミド(268mg、0.49mmol)の溶液に、K2CO3(135mg、0.98mmol)を添加した。混合物を、窒素雰囲気下で、80℃で2時間撹拌した。混合物をEA(80mL)で抽出した。有機相を水(10mL)およびブライン(10mL)で洗浄した。有機層を乾燥させ(Na2SO4)、減圧下で濾過および濃縮した。残渣を、シリカゲルカラムクロマトグラフィーによって精製して、(2S,4R)-4-ヒドロキシ-1-((S)-3-メチル-2-(1-オキソイソインドリン-2-イル)ブタノイル)-N-(4-(4-メチルチアゾール-5-イル)-2-((17-(4-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)-1H-ピラゾール-1-イル)-3,6,9,12,15-ペンタオキサヘプタデシル)オキシ)ベンジル)ピロリジン-2-カルボキサミドおよび(1-(17-(2-(((2S,4R)-4-ヒドロキシ-1-((S)-3-メチル-2-(1-オキソイソインドリン-2-イル)ブタノイル)ピロリジン-2-カルボキサミド)メチル)-5-(4-メチルチアゾール-5-イル)フェノキシ)-3,6,9,12,15-ペンタオキサヘプタデシル)-1H-ピラゾール-4-イル)ボロン酸(270mg、0.27mmol、収率56%)の混合物を得た。 To a solution of 17-(4-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)-1H-pyrazol-1-yl)-3,6,9,12,15-pentaoxaheptadecyl 4-methylbenzenesulfonate (0.3 g, 0.49 mmol) and (2S,4R)-4-hydroxy-N-(2-hydroxy-4-(4-methylthiazol-5-yl)benzyl)-1-((S)-3-methyl-2-(1-oxoisoindolin-2-yl)butanoyl)pyrrolidine-2-carboxamide (268 mg, 0.49 mmol) in DMF (5 mL) was added K2CO3 (135 mg, 0.98 mmol). The mixture was stirred at 80 °C for 2 h under nitrogen atmosphere. The mixture was extracted with EA (80 mL). The organic phase was washed with water (10 mL) and brine (10 mL). The organic layer was dried (Na2SO4), filtered and concentrated under reduced pressure. The residue was purified by silica gel column chromatography to give (2S,4R)-4-hydroxy-1-((S)-3-methyl-2-(1-oxoisoindolin-2-yl)butanoyl)-N-(4-(4-methylthiazol-5-yl)-2-((17-(4-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)-1H-pyrazol-1-yl)-3,6,9,12,15-pentaoxaheptadecyl)oxy)benzyl)pi A mixture of pyrrolidine-2-carboxamide and (1-(17-(2-(((2S,4R)-4-hydroxy-1-((S)-3-methyl-2-(1-oxoisoindolin-2-yl)butanoyl)pyrrolidine-2-carboxamide)methyl)-5-(4-methylthiazol-5-yl)phenoxy)-3,6,9,12,15-pentaoxaheptadecyl)-1H-pyrazol-4-yl)boronic acid (270 mg, 0.27 mmol, 56% yield) was obtained.
工程4

ジオキサン(20mL)および水(2mL)中の(2S,4R)-4-ヒドロキシ-1-((S)-3-メチル-2-(1-オキソイソインドリン-2-イル)ブタノイル)-N-(4-(4-メチルチアゾール-5-イル)-2-((17-(4-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)-1H-ピラゾール-1-イル)-3,6,9,12,15-ペンタオキサヘプタデシル)オキシ)ベンジル)ピロリジン-2-カルボキサミドおよび(1-(17-(2-(((2S,4R)-4-ヒドロキシ-1-((S)-3-メチル-2-(1-オキソイソインドリン-2-イル)ブタノイル)ピロリジン-2-カルボキサミド)メチル)-5-(4-メチルチアゾール-5-イル)フェノキシ)-3,6,9,12,15-ペンタオキサヘプタデシル)-1H-ピラゾール-4-イル)ボロン酸(270mg、0.27mmol)の混合物、および5-ブロモ-6-クロロピリダジン-3-アミン(85mg、0.41mmol)の溶液に、炭酸セシウム(220mg、0.68mmol)、PdCl2(dppf)(40mg、0.054mmol)、およびt-Bu3PHBF4(31mg、0.11mmol)を添加した。溶液を、窒素雰囲気下で、100℃で4時間撹拌した。混合物を減圧下で濃縮した。残渣を、シリカゲルカラムクロマトグラフィーによって精製して、(2S,4R)-N-(2-((17-(4-(3-アミノ-6-クロロピリダジン-4-イル)-1H-ピラゾール-1-イル)-3,6,9,12,15-ペンタオキサヘプタデシル)オキシ)-4-(4-メチルチアゾール-5-イル)ベンジル)-4-ヒドロキシ-1-((S)-3-メチル-2-(1-オキソイソインドリン-2-イル)ブタノイル)ピロリジン-2-カルボキサミド(175mg、0.18mmol、収率67%)を得た。 (2S,4R)-4-hydroxy-1-((S)-3-methyl-2-(1-oxoisoindolin-2-yl)butanoyl)-N-(4-(4-methylthiazol-5-yl)-2-((17-(4-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)-1H-pyrazol-1-yl)-3,6,9,12,15-pentaoxaheptadecyl)oxy)benzyl)pyrrolidine-2-carboxamide and (1-(17-(2-(((2S,4R)-4-hydroxy-1-((S)-3-methyl-2-(1-oxoisoindolin-2-yl)butanoyl)-N-(4-(4-methylthiazol-5-yl)-2-((17-(4-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)-1H-pyrazol-1-yl)-3,6,9,12,15-pentaoxaheptadecyl)oxy)benzyl)pyrrolidine-2-carboxamide in dioxane (20 mL) and water (2 mL). To a mixture of (xoisoindolin-2-yl)butanoyl)pyrrolidine-2-carboxamido)methyl)-5-(4-methylthiazol-5-yl)phenoxy)-3,6,9,12,15-pentaoxaheptadecyl)-1H-pyrazol-4-yl)boronic acid (270 mg, 0.27 mmol) and a solution of 5-bromo-6-chloropyridazin-3-amine (85 mg, 0.41 mmol) was added cesium carbonate (220 mg, 0.68 mmol), PdCl2(dppf) (40 mg, 0.054 mmol), and t-Bu3PHBF4 (31 mg, 0.11 mmol). The solution was stirred at 100° C. for 4 h under nitrogen atmosphere. The mixture was concentrated under reduced pressure. The residue was purified by silica gel column chromatography to obtain (2S,4R)-N-(2-((17-(4-(3-amino-6-chloropyridazin-4-yl)-1H-pyrazol-1-yl)-3,6,9,12,15-pentaoxaheptadecyl)oxy)-4-(4-methylthiazol-5-yl)benzyl)-4-hydroxy-1-((S)-3-methyl-2-(1-oxoisoindolin-2-yl)butanoyl)pyrrolidine-2-carboxamide (175 mg, 0.18 mmol, 67% yield).
工程5

ジオキサン(15mL)および水(1.5mL)中の(2S,4R)-N-(2-((17-(4-(3-アミノ-6-クロロピリダジン-4-イル)-1H-ピラゾール-1-イル)-3,6,9,12,15-ペンタオキサヘプタデシル)オキシ)-4-(4-メチルチアゾール-5-イル)ベンジル)-4-ヒドロキシ-1-((S)-3-メチル-2-(1-オキソイソインドリン-2-イル)ブタノイル)ピロリジン-2-カルボキサミド(175mg、0.18mmol)および(2-ヒドロキシフェニル)ボロン酸(32mg、0.23mmol)の溶液に、炭酸セシウム(176mg、0.54mmol)、PdCl2(dppf)(53mg、0.072mmol)、およびt-Bu3PHBF4(42mg、0.144mmol)を添加した。溶液を、窒素雰囲気下で、100℃で4時間撹拌した。混合物を減圧下で濃縮した。残渣を、シリカゲルカラムクロマトグラフィーによって精製して、(2S,4R)-N-(2-((17-(4-(3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル)-1H-ピラゾール-1-イル)-3,6,9,12,15-ペンタオキサヘプタデシル)オキシ)-4-(4-メチルチアゾール-5-イル)ベンジル)-4-ヒドロキシ-1-((S)-3-メチル-2-(1-オキソイソインドリン-2-イル)ブタノイル)ピロリジン-2-カルボキサミド(24mg、0.023mmol、収率13%)を得た。 (2S,4R)-N-(2-((17-(4-(3-amino-6-chloropyridazin-4-yl)-1H-pyrazol-1-yl)-3,6,9,12,15-pentaoxaheptadecyl)oxy)-4-(4-methylthiazol-5-yl)benzyl)-4-hydroxy-1-((S)-3-methyl-2-(1-oxoisoin) in dioxane (15 mL) and water (1.5 mL). To a solution of (doline-2-yl)butanoyl)pyrrolidine-2-carboxamide (175 mg, 0.18 mmol) and (2-hydroxyphenyl)boronic acid (32 mg, 0.23 mmol) was added cesium carbonate (176 mg, 0.54 mmol), PdCl2(dppf) (53 mg, 0.072 mmol), and t-Bu3PHBF4 (42 mg, 0.144 mmol). The solution was stirred at 100° C. under nitrogen atmosphere for 4 h. The mixture was concentrated under reduced pressure. The residue was purified by silica gel column chromatography to obtain (2S,4R)-N-(2-((17-(4-(3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl)-1H-pyrazol-1-yl)-3,6,9,12,15-pentaoxaheptadecyl)oxy)-4-(4-methylthiazol-5-yl)benzyl)-4-hydroxy-1-((S)-3-methyl-2-(1-oxoisoindolin-2-yl)butanoyl)pyrrolidine-2-carboxamide (24 mg, 0.023 mmol, 13% yield).
1HNMR(400MHz、MeOD):δ8.84(s、1H)、8.28(s、1H)、8.05(d、J=12.4Hz、2H)、7.38~7.81(m、6H)、7.36~7.37(m、1H)、6.98~7.00(m、2H)、6.92(d、J=7.6Hz、2H)、4.37~4.59(m、9H)、4.15~4.16(m、2H)、3.83~3.96(m、6H)、3.46~3.56(m、16H)、2.46(s、3H)、2.44~2.45(m、1H)、2.21~2.22(m、1H)、2.09~2.10(m、1H)、1.03(d、J=6.4Hz、3H)、0.82(d、J=6.4Hz、3H)。 1H NMR (400 MHz, MeOD): δ 8.84 (s, 1H), 8.28 (s, 1H), 8.05 (d, J = 12.4 Hz, 2H), 7.38-7.81 (m, 6H), 7.36-7.37 (m, 1H), 6.98-7.00 (m, 2H), 6.92 (d, J=7.6 Hz, 2H), 4.37-4. 59 (m, 9H), 4 . 15-4.16 (m, 2H), 3.83-3.96 (m, 6H), 3.46-3.56 (m, 16H), 2.46 (s, 3H), 2.44 ~2.45 (m, 1H), 2.21-2.22 (m, 1H), 2.09-2.10 (m, 1H), 1.03 (d, J = 6.4 Hz, 3H) , 0.82 (d, J = 6.4 Hz, 3H).
例示的な化合物20について記載されるものと類似の手順を使用して、例示的な化合物12、13、および21を調製した。 Exemplary compounds 12, 13, and 21 were prepared using procedures similar to those described for exemplary compound 20.
例示的な化合物14の例示的な合成
工程1

250mLの丸底フラスコに、ジオキサン/H2O(1:1)(60mL)中の4-(テトラメチル-1,3,2-ジオキサボロラン-2-イル)-1H-ピラゾール(1.5g、7.73mmol、1.00当量)の溶液、4-ブロモ-6-クロロピリダジン-3-アミン(1.7g、8.16mmol、1.20当量)、Pd(PPh3)4(800mg、0.69mmol、0.10当量)、炭酸カリウム(2.9g、20.98mmol、3.00当量)を入れた。得られた溶液を一晩、油浴中、100℃で撹拌した。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。これにより、1.0g(66%)の6-クロロ-4-(1H-ピラゾール-4-イル)ピリダジン-3-アミンを白色の固体として得た。 A 250 mL round-bottom flask was charged with a solution of 4-(tetramethyl-1,3,2-dioxaborolan-2-yl)-1H-pyrazole (1.5 g, 7.73 mmol, 1.00 equiv.), 4-bromo-6-chloropyridazin-3-amine (1.7 g, 8.16 mmol, 1.20 equiv.), Pd(PPh3)4 (800 mg, 0.69 mmol, 0.10 equiv.), and potassium carbonate (2.9 g, 20.98 mmol, 3.00 equiv.) in dioxane/H2O (1:1) (60 mL). The resulting solution was stirred overnight at 100 °C in an oil bath. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (10:1). This yielded 1.0 g (66%) of 6-chloro-4-(1H-pyrazol-4-yl)pyridazin-3-amine as a white solid.
工程2

10mLの密封チューブに、ジオキサン(4mL)中の6-クロロ-4-(1H-ピラゾール-4-イル)ピリダジン-3-アミン(390mg、1.99mmol、1.00当量)の溶液、[2-(メトキシメトキシ)フェニル]ボロン酸(546mg、3.00mmol、1.50当量)、Pd(PPh3)4(300mg、0.26mmol、0.20当量)、水(2mL)中の炭酸カリウム(552mg、3.99mmol、2.00当量)の溶液を入れた。得られた溶液を、油浴中で、100℃で12時間撹拌した。得られた混合物を真空下で濃縮した。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。これにより、120mg(20%)の6-[2-(メトキシメトキシ)フェニル]-4-(1H-ピラゾール-4-イル)ピリダジン-3-アミンを黄色の固体として得た。 A 10 mL sealed tube was charged with a solution of 6-chloro-4-(1H-pyrazol-4-yl)pyridazin-3-amine (390 mg, 1.99 mmol, 1.00 equiv.) in dioxane (4 mL), [2-(methoxymethoxy)phenyl]boronic acid (546 mg, 3.00 mmol, 1.50 equiv.), Pd(PPh3)4 (300 mg, 0.26 mmol, 0.20 equiv.), and a solution of potassium carbonate (552 mg, 3.99 mmol, 2.00 equiv.) in water (2 mL). The resulting solution was stirred in an oil bath at 100 °C for 12 h. The resulting mixture was concentrated under vacuum. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (10:1). This gave 120 mg (20%) of 6-[2-(methoxymethoxy)phenyl]-4-(1H-pyrazol-4-yl)pyridazin-3-amine as a yellow solid.
工程3

50mLの丸底フラスコに、N,N-ジメチルホルムアミド(10mL)中の6-[2-(メトキシメトキシ)フェニル]-4-(1H-ピラゾール-4-イル)ピリダジン-3-アミン(100mg、0.34mmol、1.00当量)の溶液、2-[2-(2-[[(4-メチルベンゼン)スルホニル]オキシ]エトキシ)エトキシ]エタン-1-オール(100mg、0.33mmol、1.00当量)、炭酸カリウム(91mg、0.66mmol、2.00当量)を入れた。得られた溶液を、油浴中で、70℃で12時間撹拌した。得られた溶液を酢酸エチル(20mL×3)で抽出し、有機層を合わせて、真空下で濃縮した。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。これにより、100mg(69%)の2-[2-[2-(4-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-1H-ピラゾール-1-イル)エトキシ]エトキシ]エタン-1-オールを黄色の油として得た。 A 50 mL round bottom flask was charged with a solution of 6-[2-(methoxymethoxy)phenyl]-4-(1H-pyrazol-4-yl)pyridazin-3-amine (100 mg, 0.34 mmol, 1.00 equiv.) in N,N-dimethylformamide (10 mL), 2-[2-(2-[[(4-methylbenzene)sulfonyl]oxy]ethoxy)ethoxy]ethan-1-ol (100 mg, 0.33 mmol, 1.00 equiv.), potassium carbonate (91 mg, 0.66 mmol, 2.00 equiv.). The resulting solution was stirred in an oil bath at 70°C for 12 hours. The resulting solution was extracted with ethyl acetate (20 mL x 3), the organic layers were combined and concentrated under vacuum. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (10:1). This gave 100 mg (69%) of 2-[2-[2-(4-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-1H-pyrazol-1-yl)ethoxy]ethoxy]ethan-1-ol as a yellow oil.
工程4

100mLの丸底フラスコに、ジクロロメタン(20mL)中の2-[2-[2-(4-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-1H-ピラゾール-1-イル)エトキシ]エトキシ]エタン-1-オール(100mg、0.23mmol、1.00当量)の溶液、4-トルエンスルホニルクロリド(66.0mg、0.35mmol、1.50当量)、トリエチルアミン(47mg、0.46mmol、2.00当量)、4-ジメチルアミノピリジン(10mg、0.08mmol、0.30当量)を入れた。得られた溶液を室温で16時間撹拌した。得られた溶液を酢酸エチル(20mL×3)で抽出し、有機層を合わせて、真空下で濃縮した。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。これにより、100mg(74%)の2-[2-[2-(4-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-1H-ピラゾール-1-イル)エトキシ]エトキシ]エチル4-メチルベンゼン-1-スルホネートを黄色の油として得た。 A 100 mL round bottom flask was charged with a solution of 2-[2-[2-(4-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-1H-pyrazol-1-yl)ethoxy]ethoxy]ethan-1-ol (100 mg, 0.23 mmol, 1.00 equiv.) in dichloromethane (20 mL), 4-toluenesulfonyl chloride (66.0 mg, 0.35 mmol, 1.50 equiv.), triethylamine (47 mg, 0.46 mmol, 2.00 equiv.), 4-dimethylaminopyridine (10 mg, 0.08 mmol, 0.30 equiv.). The resulting solution was stirred at room temperature for 16 hours. The resulting solution was extracted with ethyl acetate (20 mL x 3), the organic layers were combined and concentrated under vacuum. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (10:1). This gave 100 mg (74%) of 2-[2-[2-(4-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-1H-pyrazol-1-yl)ethoxy]ethoxy]ethyl 4-methylbenzene-1-sulfonate as a yellow oil.
工程5

50mLの丸底フラスコに、N,N-ジメチルホルムアミド(5mL)中の2-[2-[2-(4-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-1H-ピラゾール-1-イル)エトキシ]エトキシ]エチル4-メチルベンゼン-1-スルホネート(120mg、0.2mmol、1.00当量)の溶液、(2S,4R)-4-ヒドロキシ-1-[2-(3-ヒドロキシ-1,2-オキサゾール-5-イル)-3-メチルブタノイル]-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]ピロリジン-2-カルボキサミド(130mg、0.2mmol、1.00当量)[Qian,YらによってWO2017/030814に説明されるように調製]、炭酸カリウム(100mg、0.4mmol、2.00当量)を入れた。得られた溶液を70℃で12時間撹拌した。得られた溶液を酢酸エチル(20mL×3)で抽出し、有機層を合わせ、真空下で濃縮した。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。これにより、90mgの(2S,4R)-1-[2-[3-(2-[2-[2-(4-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-1H-ピラゾール-1-イル)エトキシ]エトキシ]エトキシ)-1,2-オキサゾール-5-イル]-3-メチルブタノイル]-4-ヒドロキシ-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]ピロリジン-2-カルボキサミドを無色の油として得た。 In a 50 mL round-bottom flask, add a solution of 2-[2-[2-(4-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-1H-pyrazol-1-yl)ethoxy]ethoxy]ethyl 4-methylbenzene-1-sulfonate (120 mg, 0.2 mmol, 1.00 equiv.) in N,N-dimethylformamide (5 mL), (2S,4R)-4-hydroxy-1-[2-(3- Hydroxy-1,2-oxazol-5-yl)-3-methylbutanoyl]-N-[[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]pyrrolidine-2-carboxamide (130 mg, 0.2 mmol, 1.00 equiv.) [prepared as described by Qian, Y et al. in WO2017/030814], potassium carbonate (100 mg, 0.4 mmol, 2.00 equiv.) were added. The resulting solution was stirred at 70° C. for 12 hours. The resulting solution was extracted with ethyl acetate (20 mL×3) and the organic layers were combined and concentrated under vacuum. The residue was applied onto a silica gel column eluting with dichloromethane/methanol (10:1). This gave 90 mg of (2S,4R)-1-[2-[3-(2-[2-[2-(4-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-1H-pyrazol-1-yl)ethoxy]ethoxy]ethoxy)-1,2-oxazol-5-yl]-3-methylbutanoyl]-4-hydroxy-N-[[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]pyrrolidine-2-carboxamide as a colorless oil.
工程6

50mLの丸底フラスコに、i-プロパノール(2mL)およびテトラヒドロフラン(2mL)中の(2S,4R)-1-[2-[3-(2-[2-[2-(4-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-1H-ピラゾール-1-イル)エトキシ]エトキシ]エトキシ)-1,2-オキサゾール-5-イル]-3-メチルブタノイル]-4-ヒドロキシ-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]ピロリジン-2-カルボキサミド(90.0mg、0.10mmol、1.00当量)の溶液を入れ、濃縮塩化水素溶液(12N、2mL)を添加した。得られた溶液を室温で1時間撹拌した。得られた混合物を真空下で濃縮した。以下の条件の分取HPLCによって、粗生成物を精製した:カラム:XBridge C18 OBD Prep Column、100Å、5μm、19mm×250mm;移動相A:水(0.1%FA)、移動相B:ACN、流速:20mL/分、勾配:8分間で32%B~41%B、254nm、保持時間:70分。これにより、56mg(65%)の(2S,4R)-1-[2-(3-[2-[2-(2-[4-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-1H-ピラゾール-1-イル]エトキシ)エトキシ]エトキシ]-1,2-オキサゾール-5-イル)-3-メチルブタノイル]-4-ヒドロキシ-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]ピロリジン-2-カルボキサミドを白色の固体として得た。 A 50 mL round bottom flask was charged with a solution of (2S,4R)-1-[2-[3-(2-[2-[2-(4-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-1H-pyrazol-1-yl)ethoxy]ethoxy]ethoxy)-1,2-oxazol-5-yl]-3-methylbutanoyl]-4-hydroxy-N-[[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]pyrrolidine-2-carboxamide (90.0 mg, 0.10 mmol, 1.00 equiv.) in i-propanol (2 mL) and tetrahydrofuran (2 mL) and concentrated hydrogen chloride solution (12 N, 2 mL) was added. The resulting solution was stirred at room temperature for 1 h. The resulting mixture was concentrated under vacuum. The crude product was purified by preparative HPLC with the following conditions: Column: XBridge C18 OBD Prep Column, 100 Å, 5 μm, 19 mm×250 mm; Mobile phase A: water (0.1% FA), Mobile phase B: ACN, Flow rate: 20 mL/min, Gradient: 32% B to 41% B in 8 min, 254 nm, Retention time: 70 min. This gave 56 mg (65%) of (2S,4R)-1-[2-(3-[2-[2-(2-[4-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-1H-pyrazol-1-yl]ethoxy)ethoxy]ethoxy]-1,2-oxazol-5-yl)-3-methylbutanoyl]-4-hydroxy-N-[[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]pyrrolidine-2-carboxamide as a white solid.
生成物を、以下の条件を用いて、キラル分取HPLCによって精製した。カラム、CHIRAL ART Cellulose-SB、2*25cm、5um;移動相、Hex--HPLCおよびエタノール--HPLC(24分間で50%エタノール--HPLCを保持);検出器、UV220/254nm。これにより、17.8mg(34%)の(2S,4R)-1-[(2S)-2-(3-[2-[2-(2-[4-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-1H-ピラゾール-1-イル]エトキシ)エトキシ]エトキシ]-1,2-オキサゾール-5-イル)-3-メチルブタノイル]-4-ヒドロキシ-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]ピロリジン-2-カルボキサミドを白色の固体[1H NMR(300MHz、CD3OD):δ8.82(d、J=11.1Hz、1H)、8.29~8.18(m、1H)、8.11~7.89(m、2H)、7.80(d、J=8.2Hz、1H)、7.48~7.41(m、1H)、7.37~7.30(m、3H)、7.30~7.19(m、1H)、6.95~6.83(m、2H)、5.89(s、1H)、4.59~4.41(m、2H)、4.40~4.30(m、4H)、4.27~4.11(m、2H)、3.86(q、J=5.3Hz、2H)、3.75~3.59(m、6H)、3.56(s、3H)、2.42(d、J=5.0Hz、3H)、2.36~2.10(m、2H)、2.04~2.01(m、J=13.1、8.1、4.7Hz、1H)、1.26(s、1H)、0.99(d、J=6.6Hz、3H)、0.83(d、J=6.8Hz、3H)]として、および24.4mg(47%)の(2S,4R)-1-[(2R)-2-(3-[2-[2-(2-[4-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-1H-ピラゾール-1-イル]エトキシ)エトキシ]エトキシ]-1,2-オキサゾール-5-イル)-3-メチルブタノイル]-4-ヒドロキシ-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]ピロリジン-2-カルボキサミド[1H NMR(300MHz、CD3OD):δ8.82(d、J=6.6Hz、1H)、8.26(dd、J=3.1、0.8Hz、1H)、8.10~7.96(m、2H)、7.86~7.75(m、1H)、7.45~7.30(m、4H)、7.25~7.22(m、1H)、6.96~6.84(m、2H)、5.88(s、1H)、4.70~4.42(m、3H)、4.41~4.33(m、3H)、4.28~4.17(m、2H)、3.87(t、J=5.0Hz、3H)、3.85~3.63(m、2H)、3.59~3.57(m、6H)、2.41(d、J=12.2Hz、3H)、2.38~2.33(m、1H)、2.25~2.12(m、1H)、2.06~2.02(m、1H)、1.26(s、1H)、0.97(dd、J=6.6、1.8Hz、3H)、0.80(dd、J=6.6、1.8Hz、3H)]を白色の固体として得た。 The product was purified by chiral preparative HPLC using the following conditions: Column, CHIRAL ART Cellulose-SB, 2*25cm, 5um; Mobile phase, Hex--HPLC and Ethanol--HPLC (50% Ethanol--HPLC held for 24min); Detector, UV 220/254nm. This gave 17.8 mg (34%) of (2S,4R)-1-[(2S)-2-(3-[2-[2-(2-[4-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-1H-pyrazol-1-yl]ethoxy)ethoxy]ethoxy]-1,2-oxazol-5-yl)-3-methylbutanoyl]-4-hydroxy-N-[[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]pyrrolidine-2-carboxamide as a white solid [1H NMR (300 MHz, CD3OD): δ 8.82 (d, J = 11.1 Hz, 1H), 8.29-8.18 (m, 1H), 8.11-7.89 (m, 2H), 7.80 (d, J = 8.2 Hz, 1H), 7.48-7.41 (m, 1H), 7.37-7.30 (m, 3H), 7.30-7.19 (m, 1H), 6.95-6.8 3 (m, 2H), 5.89 (s, 1H), 4.59-4.41 (m, 2H), 4.40-4.30 (m, 4H), 4.27-4.11 (m, 2H), 3.86 (q, J = 5.3 Hz, 2H), 3.75-3.59 (m, 6H), 3.56 (s, 3H), 2.42 (d, J = 5.0 Hz, 3H), 2.36-2.10 (m, 2H ), 2.04-2.01 (m, J = 13.1, 8.1, 4.7 Hz, 1H), 1.26 (s, 1H), 0.99 (d, J = 6.6 Hz, 3H), 0.83 (d, J = 6.8 Hz, 3H)], and 24.4 mg (47%) of (2S,4R)-1-[(2R)-2-(3-[2-[2-(2-[4-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-1H-pyrazol-1-yl]ethoxy)ethoxy]ethoxy]-1,2-oxazol-5-yl)-3-methylbutanoyl]-4-hydroxy-N-[[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]pyrrolidine-2-carboxamide [1H NMR (300 MHz, CD3OD): δ 8.82 (d, J = 6.6 Hz, 1H), 8.26 (dd, J = 3.1, 0.8 Hz, 1H), 8.10-7.96 (m, 2H), 7.86-7.75 (m, 1H), 7.45-7.30 (m, 4H), 7.25-7.22 (m, 1H), 6.96-6.84 (m, 2H), 5.88 (s, 1H), 4.70-4.42 (m, 3H), 4.41-4.33 (m, 3H), 4.28-4.17 (m , 2H), 3.87 (t, J = 5.0 Hz, 3H), 3.85-3.63 (m, 2H), 3.59-3.57 (m, 6H), 2.41 (d, J = 12.2 Hz, 3H), 2.38-2.33 (m, 1H), 2.25-2.12 (m, 1H), 2.06-2.02 (m, 1H), 1.26 (s, 1H), 0.97 (dd, J = 6.6, 1.8 Hz, 3H), 0.80 (dd, J = 6.6, 1.8 Hz, 3H)] was obtained as a white solid.
例示的な化合物16、17、18、および19を、例示的な化合物14および例示的な化合物15について上に記載される手順を使用して調製した。 Exemplary compounds 16, 17, 18, and 19 were prepared using the procedures described above for exemplary compound 14 and exemplary compound 15.
例示的な化合物22の例示的な合成
工程1
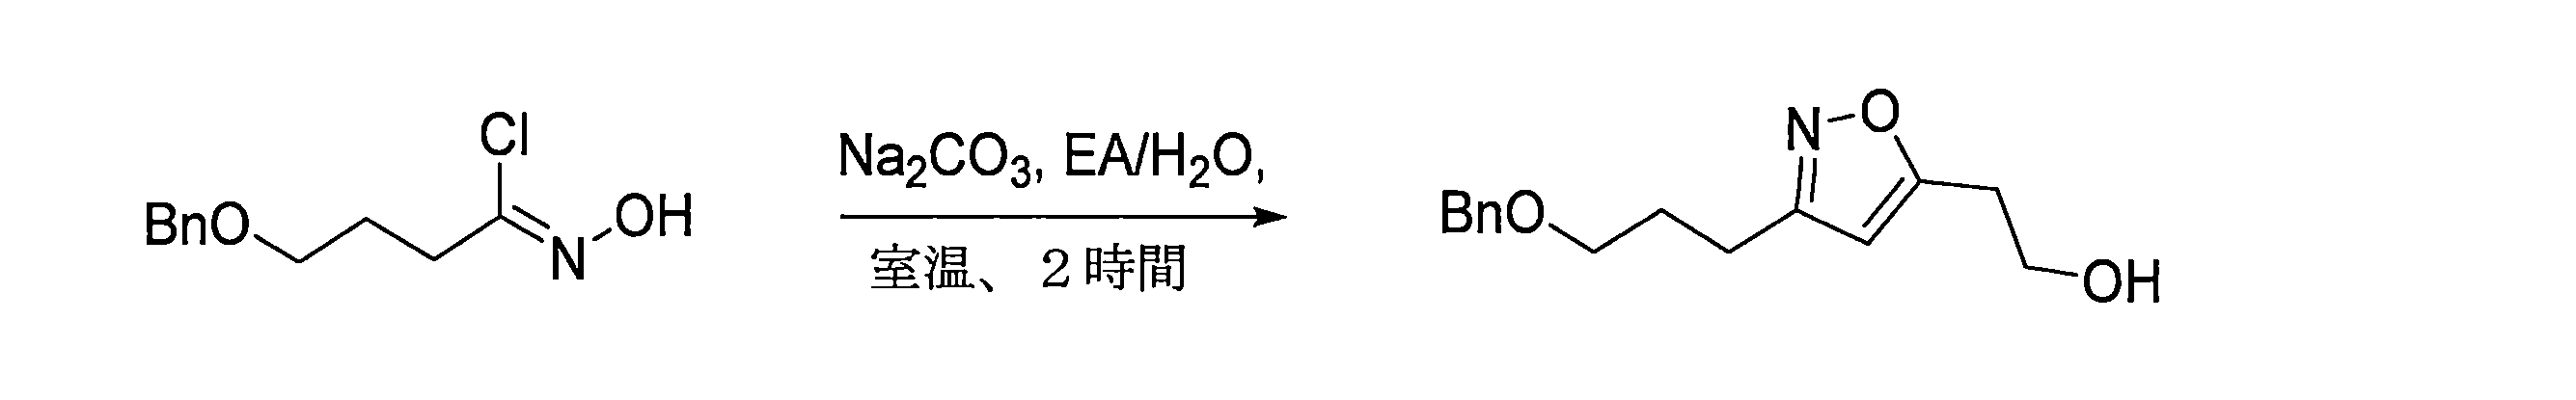
250mLの丸底フラスコに、(Z)-4-(ベンジルオキシ)-N-ヒドロキシブトカルボニミドイルクロリド(8.7g、38.21mmol、1.00当量)、ブタ-3-イン-1-オール(3.3g、47.08mmol、1.23当量)、酢酸エチル(70mL)、水(70mL)、重炭酸ナトリウム(4.0g、47.61mmol、1.25当量)を入れた。得られた溶液を25℃で2時間撹拌した。得られた溶液を酢酸エチルで抽出し、飽和塩化ナトリウム水溶液で洗浄した。混合物を、無水硫酸ナトリウム上で乾燥させ、真空下で濃縮した。残渣を、酢酸エチル/石油エーテル(1:1)で溶出するシリカゲルカラム上に適用した。これにより、5.9g(59%)の2-[3-[3-(ベンジルオキシ)プロピル]-1,2-オキサゾール-5-イル]エタン-1-オールを黄色の油として得た。 A 250 mL round bottom flask was charged with (Z)-4-(benzyloxy)-N-hydroxybutocarbonimidoyl chloride (8.7 g, 38.21 mmol, 1.00 equiv.), but-3-yn-1-ol (3.3 g, 47.08 mmol, 1.23 equiv.), ethyl acetate (70 mL), water (70 mL), and sodium bicarbonate (4.0 g, 47.61 mmol, 1.25 equiv.). The resulting solution was stirred at 25° C. for 2 h. The resulting solution was extracted with ethyl acetate and washed with saturated aqueous sodium chloride. The mixture was dried over anhydrous sodium sulfate and concentrated under vacuum. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (1:1). This gave 5.9 g (59%) of 2-[3-[3-(benzyloxy)propyl]-1,2-oxazol-5-yl]ethan-1-ol as a yellow oil.
工程2

100mLの丸底フラスコに、2-[3-[3-(ベンジルオキシ)プロピル]-1,2-オキサゾール-5-イル]エタン-1-オール(550.0mg、2.10mmol、1.00当量)、アセトン(30mL)、Cr2O3(100.0mg)、硫酸(0.25mL)、水(1mL)を入れた。得られた溶液を25℃で1時間撹拌した。得られた溶液を水で希釈した。得られた溶液を酢酸エチルで抽出し、飽和塩化ナトリウム水溶液で洗浄した。混合物を、無水硫酸ナトリウム上で乾燥させ、真空下で濃縮した。これにより、420mg(72%)の2-[3-[3-(ベンジルオキシ)プロピル]-1,2-オキサゾール-5-イル]酢酸を黄色の油として得た。 A 100 mL round bottom flask was charged with 2-[3-[3-(benzyloxy)propyl]-1,2-oxazol-5-yl]ethan-1-ol (550.0 mg, 2.10 mmol, 1.00 equiv.), acetone (30 mL), Cr2O3 (100.0 mg), sulfuric acid (0.25 mL), and water (1 mL). The resulting solution was stirred at 25°C for 1 h. The resulting solution was diluted with water. The resulting solution was extracted with ethyl acetate and washed with saturated aqueous sodium chloride. The mixture was dried over anhydrous sodium sulfate and concentrated under vacuum. This afforded 420 mg (72%) of 2-[3-[3-(benzyloxy)propyl]-1,2-oxazol-5-yl]acetic acid as a yellow oil.
工程3
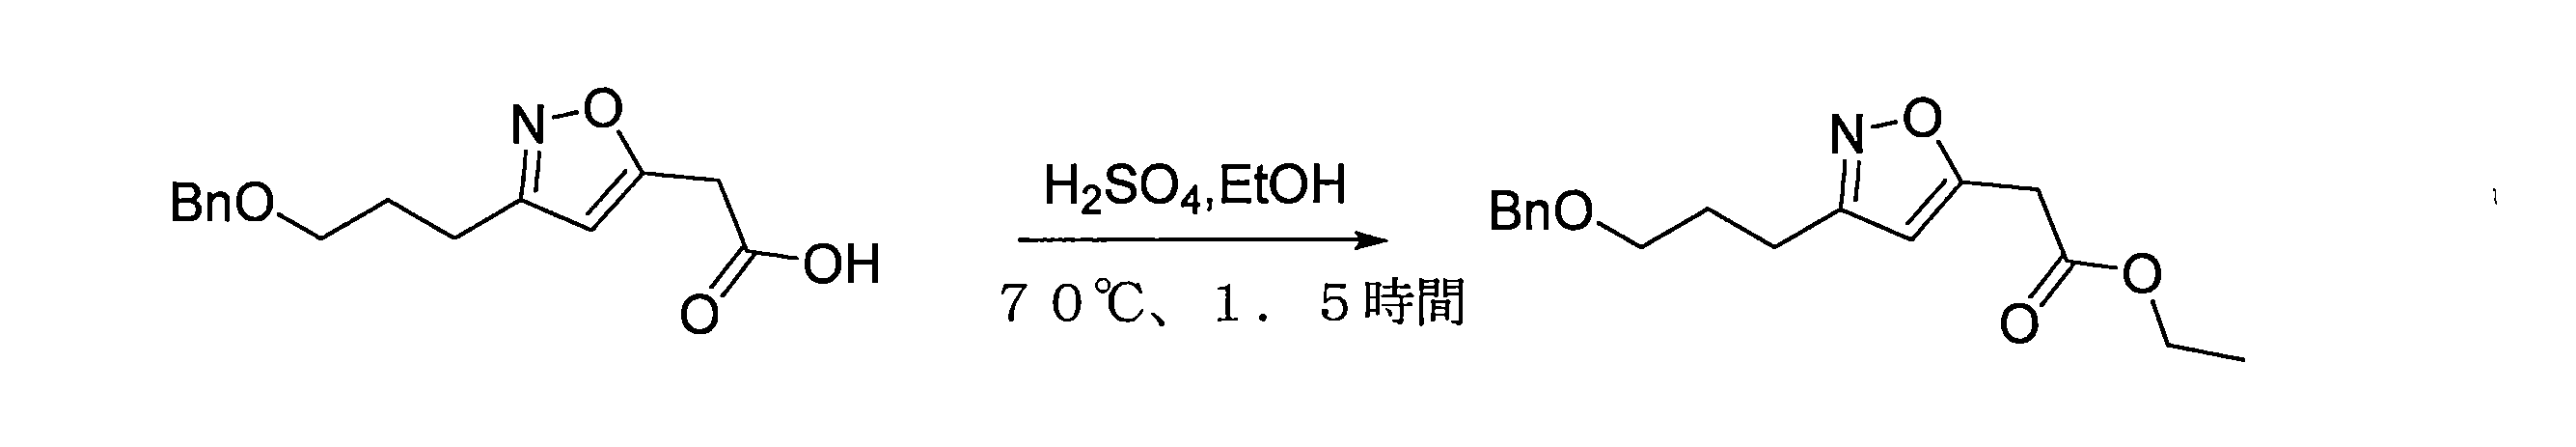
250mLの丸底フラスコに、エチル2-[3-[3-(ベンジルオキシ)プロピル]-1,2-オキサゾール-5-イル]酢酸塩(8.0g、26.37mmol、1.00当量)、エタノール(50mL)、硫酸(0.1mL)を入れた。得られた溶液を70℃で1.5時間撹拌した。得られた混合物を、真空下で濃縮した。粗生成物を、以下の条件を用いて、フラッシュ分取HPLCによって精製した。カラム、C18シリカゲル;移動相、49分以内にアセトニトリル:水=0:100がアセトニトリル:水=60:40に増加;検出器、UV220nm。これにより、4.5g(56%)のエチル2-[3-[3-(ベンジルオキシ)プロピル]-1,2-オキサゾール-5-イル]酢酸塩を淡い黄色の油として得た。 A 250 mL round bottom flask was charged with ethyl 2-[3-[3-(benzyloxy)propyl]-1,2-oxazol-5-yl]acetate (8.0 g, 26.37 mmol, 1.00 equiv), ethanol (50 mL), and sulfuric acid (0.1 mL). The resulting solution was stirred at 70° C. for 1.5 h. The resulting mixture was concentrated under vacuum. The crude product was purified by flash preparative HPLC using the following conditions: column, C18 silica gel; mobile phase, acetonitrile:water=0:100 increasing to acetonitrile:water=60:40 within 49 min; detector, UV 220 nm. This gave 4.5 g (56%) of ethyl 2-[3-[3-(benzyloxy)propyl]-1,2-oxazol-5-yl]acetate as a pale yellow oil.
工程4
窒素不活性雰囲気下でパージし、維持した250mLの三口丸底フラスコに、エチル2-[3-[3-(ベンジルオキシ)プロピル]-1,2-オキサゾール-5-イル]酢酸塩(4.5g、14.83mmol、1.00当量)およびテトラヒドロフラン(70mL)を入れた。これに続いて、テトラヒドロフラン(17.8mL)中のt-BuOK(2.0g、17.82mmol、1.20当量)の溶液を、0℃で20分間撹拌しながら滴下で添加した。これに、2-ヨードプロパン(3.01g、17.71mmol、1.19当量)を、0℃で2分間撹拌しながら滴下で添加した。得られた溶液を25℃で2時間撹拌した。次いで、水によって反応物をクエンチした。得られた溶液を酢酸エチルで抽出し、飽和塩化ナトリウム水溶液で洗浄した。混合物を、無水硫酸ナトリウム上で乾燥させ、真空下で濃縮した。これにより、4.3g(84%)のエチル2-[3-[3-(ベンジルオキシ)プロピル]-1,2-オキサゾール-5-イル]-3-メチルブタノエートをオレンジ色の油として得た。 A 250 mL three-neck round bottom flask purged and maintained under nitrogen inert atmosphere was charged with ethyl 2-[3-[3-(benzyloxy)propyl]-1,2-oxazol-5-yl]acetate (4.5 g, 14.83 mmol, 1.00 equiv.) and tetrahydrofuran (70 mL). Following this, a solution of t-BuOK (2.0 g, 17.82 mmol, 1.20 equiv.) in tetrahydrofuran (17.8 mL) was added dropwise with stirring for 20 minutes at 0°C. To this, 2-iodopropane (3.01 g, 17.71 mmol, 1.19 equiv.) was added dropwise with stirring for 2 minutes at 0°C. The resulting solution was stirred at 25°C for 2 hours. The reaction was then quenched by water. The resulting solution was extracted with ethyl acetate and washed with saturated aqueous sodium chloride. The mixture was dried over anhydrous sodium sulfate and concentrated under vacuum. This gave 4.3 g (84%) of ethyl 2-[3-[3-(benzyloxy)propyl]-1,2-oxazol-5-yl]-3-methylbutanoate as an orange oil.
工程5

窒素不活性雰囲気下でパージし、維持した250mLの三口丸底フラスコに、エチル2-[3-[3-(ベンジルオキシ)プロピル]-1,2-オキサゾール-5-イル]-3-メチルブタノエート(4.2g、12.16mmol、1.00当量)およびジクロロメタン(100mL)を入れた。これに続いて、ジクロロメタン(20.7mL)中のBBr3(5.17g、20.64mmol、1.70当量)の溶液を、-78℃で30分間撹拌しながら滴下で添加した。得られた溶液を-78℃で2時間撹拌した。次いで、反応物をクエンチした。得られた溶液をジクロロメタンで抽出し、飽和塩化ナトリウム水溶液で洗浄した。混合物を、無水硫酸ナトリウム上で乾燥させ、真空下で濃縮した。粗生成物を、以下の条件を用いて、フラッシュ分取HPLCによって精製した。カラム、C18シリカゲル;移動相、25分以内にアセトニトリル:水=0:100がアセトニトリル:水=23:76に増加;検出器、UV220nm。これにより、2.6g(84%)のエチル2-[3-(3-ヒドロキシプロピル)-1,2-オキサゾール-5-イル]-3-メチルブタノエートをオレンジ色の油として得た。 A 250 mL three-neck round bottom flask purged and maintained under a nitrogen inert atmosphere was charged with ethyl 2-[3-[3-(benzyloxy)propyl]-1,2-oxazol-5-yl]-3-methylbutanoate (4.2 g, 12.16 mmol, 1.00 equiv.) and dichloromethane (100 mL). Following this, a solution of BBr3 (5.17 g, 20.64 mmol, 1.70 equiv.) in dichloromethane (20.7 mL) was added dropwise with stirring at −78° C. for 30 min. The resulting solution was stirred at −78° C. for 2 h. The reaction was then quenched. The resulting solution was extracted with dichloromethane and washed with saturated aqueous sodium chloride. The mixture was dried over anhydrous sodium sulfate and concentrated under vacuum. The crude product was purified by flash preparative HPLC using the following conditions: Column: C18 silica gel; mobile phase: acetonitrile:water = 0:100 increasing to acetonitrile:water = 23:76 within 25 minutes; detector: UV 220 nm. This gave 2.6 g (84%) of ethyl 2-[3-(3-hydroxypropyl)-1,2-oxazol-5-yl]-3-methylbutanoate as an orange oil.
工程6

100mLの丸底フラスコに、エチル2-[3-(3-ヒドロキシプロピル)-1,2-オキサゾール-5-イル]-3-メチルブタノエート(1.2g、4.70mmol、1.00当量)、エタノール(20mL)、水(10mL)、水酸化ナトリウム(1.9g、47.50mmol、10.0当量)を入れた。得られた溶液を室温で一晩撹拌した。溶液のpH値を、塩化水素(2M)で6に調整した。得られた溶液を酢酸エチルで抽出し、飽和塩化ナトリウムで洗浄した。混合物を、無水硫酸ナトリウム上で乾燥させ、真空下で濃縮した。これにより、800mg(75%)の2-[3-(3-ヒドロキシプロピル)-1,2-オキサゾール-5-イル]-3-メチルブタン酸を黄色の油として得た。 A 100 mL round bottom flask was charged with ethyl 2-[3-(3-hydroxypropyl)-1,2-oxazol-5-yl]-3-methylbutanoate (1.2 g, 4.70 mmol, 1.00 equiv.), ethanol (20 mL), water (10 mL), and sodium hydroxide (1.9 g, 47.50 mmol, 10.0 equiv.). The resulting solution was stirred at room temperature overnight. The pH value of the solution was adjusted to 6 with hydrogen chloride (2 M). The resulting solution was extracted with ethyl acetate and washed with saturated sodium chloride. The mixture was dried over anhydrous sodium sulfate and concentrated under vacuum. This gave 800 mg (75%) of 2-[3-(3-hydroxypropyl)-1,2-oxazol-5-yl]-3-methylbutanoic acid as a yellow oil.
工程7
100mLの丸底フラスコに、2-[3-(3-ヒドロキシプロピル)-1,2-オキサゾール-5-イル]-3-メチルブタン酸(800mg、3.52mmol、1.00当量)、(2S,4R)-4-ヒドロキシ-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]ピロリジン-2-カルボキサミド塩酸塩(1.24g、3.50mmol、1.00当量)、N,N-ジメチルホルムアミド(15mL)、N-エチル-N-イソプロピルプロパン-2-アミン(1.82g、14.08mmol、4.00当量)、T3P(1.77g、1.20当量)を入れた。得られた溶液を室温で2時間撹拌した。次いで、反応物を水でクエンチした。得られた溶液を酢酸エチルで抽出し、飽和塩化ナトリウムで洗浄した。混合物を、無水硫酸ナトリウム上で乾燥させ、真空下で濃縮した。残渣を、ジクロロメタン/メタノール(5:1)で溶出するシリカゲルカラム上に適用した。収集した画分をまとめ、真空下で濃縮した。これにより、870mg(53%)の(2S,4R)-4-ヒドロキシ-1-[[3-(3-ヒドロキシプロピル)-1,2-オキサゾール-5-イル]カルボニル]-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]ピロリジン-2-カルボキサミドを黄色の固体として得た。 A 100 mL round bottom flask was charged with 2-[3-(3-hydroxypropyl)-1,2-oxazol-5-yl]-3-methylbutanoic acid (800 mg, 3.52 mmol, 1.00 equiv.), (2S,4R)-4-hydroxy-N-[[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]pyrrolidine-2-carboxamide hydrochloride (1.24 g, 3.50 mmol, 1.00 equiv.), N,N-dimethylformamide (15 mL), N-ethyl-N-isopropylpropan-2-amine (1.82 g, 14.08 mmol, 4.00 equiv.), and T3P (1.77 g, 1.20 equiv.). The resulting solution was stirred at room temperature for 2 hours. The reaction was then quenched with water. The resulting solution was extracted with ethyl acetate and washed with saturated sodium chloride. The mixture was dried over anhydrous sodium sulfate and concentrated under vacuum. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (5:1). The collected fractions were combined and concentrated under vacuum. This gave 870 mg (53%) of (2S,4R)-4-hydroxy-1-[[3-(3-hydroxypropyl)-1,2-oxazol-5-yl]carbonyl]-N-[[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]pyrrolidine-2-carboxamide as a yellow solid.
工程8

50mLの丸底フラスコに、(2S,4R)-4-ヒドロキシ-1-[2-[3-(3-ヒドロキシプロピル)-1,2-オキサゾール-5-イル]-3-メチルブタノイル]-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]ピロリジン-2-カルボキサミド(870mg、1.65mmol、1.00当量)、ジクロロメタン(10mL)、4-メチルベンゼン-1-スルホニルクロリド(377mg、1.98mmol、1.20当量)、トリエチルアミン(334.0mg、3.30mmol、2.00当量)、4-ジメチルアミノピリジン(40mg、0.33mmol、0.20当量)を入れた。得られた溶液を室温で4時間撹拌した。得られた溶液をジクロロメタンで抽出し、水および飽和塩化ナトリウムで洗浄した。混合物を、無水硫酸ナトリウム上で乾燥させ、真空下で濃縮した。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。収集した画分をまとめ、真空下で濃縮した。これにより、600mg(53%)の3-(5-[1-[(2S,4R)-4-ヒドロキシ-2-([[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]カルバモイル)ピロリジン-1-イル]-3-メチル-1-オキソブタン-2-イル]-1,2-オキサゾール-3-イル)プロピル4-メチルベンゼン-1-スルホネートを黄色の固体として得た。 A 50 mL round-bottom flask was charged with (2S,4R)-4-hydroxy-1-[2-[3-(3-hydroxypropyl)-1,2-oxazol-5-yl]-3-methylbutanoyl]-N-[[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]pyrrolidine-2-carboxamide (870 mg, 1.65 mmol, 1.00 equiv.), dichloromethane (10 mL), 4-methylbenzene-1-sulfonyl chloride (377 mg, 1.98 mmol, 1.20 equiv.), triethylamine (334.0 mg, 3.30 mmol, 2.00 equiv.), and 4-dimethylaminopyridine (40 mg, 0.33 mmol, 0.20 equiv.). The resulting solution was stirred at room temperature for 4 hours. The resulting solution was extracted with dichloromethane and washed with water and saturated sodium chloride. The mixture was dried over anhydrous sodium sulfate and concentrated under vacuum. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (10:1). The collected fractions were combined and concentrated under vacuum. This gave 600 mg (53%) of 3-(5-[1-[(2S,4R)-4-hydroxy-2-([[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]carbamoyl)pyrrolidin-1-yl]-3-methyl-1-oxobutan-2-yl]-1,2-oxazol-3-yl)propyl 4-methylbenzene-1-sulfonate as a yellow solid.
工程9

50mLの丸底フラスコに、2-(2-ヒドロキシエトキシ)エタン-1-オール(273.0mg、2.57mmol、5.00当量)およびN,N-ジメチルホルムアミド(5mL)を入れた。水素化ナトリウム(41.0mg、1.71mmol、2.00当量)を0℃下で添加し、続いて、20分後に、3-(5-[1-[(4S)-4-ヒドロキシ-1-([[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]カルバモイル)ピロリジン-2-イル]-3-メチル-1-オキソブタン-2-イル]-1,2-オキサゾール-3-イル)プロピル4-メチルベンゼン-1-スルホネート(350.0mg、0.51mmol、1.00当量)を添加した。得られた溶液を室温で4時間撹拌した。反応混合物を水で希釈し、DCMで抽出した。有機相を硫酸ナトリウムで乾燥させて、濃縮し、残渣をフラッシュクロマトグラフィーに供した。これにより、160mg(51%)の(4S)-4-ヒドロキシ-2-[2-(3-[3-[2-(2-ヒドロキシエトキシ)エトキシ]プロピル]-1,2-オキサゾール-5-イル)-3-メチルブタノイル]-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]ピロリジン-1-カルボキサミドを黄色の油として得た。 A 50 mL round bottom flask was charged with 2-(2-hydroxyethoxy)ethan-1-ol (273.0 mg, 2.57 mmol, 5.00 equiv.) and N,N-dimethylformamide (5 mL). Sodium hydride (41.0 mg, 1.71 mmol, 2.00 equiv.) was added at 0°C, followed after 20 min by the addition of 3-(5-[1-[(4S)-4-hydroxy-1-([[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]carbamoyl)pyrrolidin-2-yl]-3-methyl-1-oxobutan-2-yl]-1,2-oxazol-3-yl)propyl 4-methylbenzene-1-sulfonate (350.0 mg, 0.51 mmol, 1.00 equiv.). The resulting solution was stirred at room temperature for 4 h. The reaction mixture was diluted with water and extracted with DCM. The organic phase was dried over sodium sulfate, concentrated, and the residue was subjected to flash chromatography to give 160 mg (51%) of (4S)-4-hydroxy-2-[2-(3-[3-[2-(2-hydroxyethoxy)ethoxy]propyl]-1,2-oxazol-5-yl)-3-methylbutanoyl]-N-[[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]pyrrolidine-1-carboxamide as a yellow oil.
工程10

50mLの丸底フラスコに、(2S,4R)-4-ヒドロキシ-1-[2-(3-[3-[2-(2-ヒドロキシエトキシ)エトキシ]プロピル]-1,2-オキサゾール-5-イル)-3-メチルブタノイル]-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]ピロリジン-2-カルボキサミド(160mg、0.26mmol、1.00当量)、ジクロロメタン(5mL)、4-メチルベンゼン-1-スルホニルクロリド(59mg、0.31mmol、1.20当量)、トリエチルアミン(53mg、0.52mmol、2.00当量)、4-ジメチルアミノピリジン(6mg、0.05mmol、0.20当量)を入れた。得られた溶液を室温で5時間撹拌した。得られた溶液をジクロロメタンで抽出し、水で洗浄した。混合物を、無水硫酸ナトリウム上で乾燥させ、真空下で濃縮した。残渣を、ジクロロメタン/メタノール(12:1)で溶出するシリカゲルカラム上に適用した。収集した画分をまとめ、真空下で濃縮した。これにより、72mg(36%)の2-[2-[3-(5-[1-[(2S,4R)-4-ヒドロキシ-2-([[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]カルバモイル)ピロリジン-1-イル]-3-メチル-1-オキソブタン-2-イル]-1,2-オキサゾール-3-イル)プロポキシ]エトキシ]エチル4-メチルベンゼン-1-スルホネートを黄色の油として得た。 A 50 mL round-bottom flask was charged with (2S,4R)-4-hydroxy-1-[2-(3-[3-[2-(2-hydroxyethoxy)ethoxy]propyl]-1,2-oxazol-5-yl)-3-methylbutanoyl]-N-[[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]pyrrolidine-2-carboxamide (160 mg, 0.26 mmol, 1.00 equiv.), dichloromethane (5 mL), 4-methylbenzene-1-sulfonyl chloride (59 mg, 0.31 mmol, 1.20 equiv.), triethylamine (53 mg, 0.52 mmol, 2.00 equiv.), and 4-dimethylaminopyridine (6 mg, 0.05 mmol, 0.20 equiv.). The resulting solution was stirred at room temperature for 5 hours. The resulting solution was extracted with dichloromethane and washed with water. The mixture was dried over anhydrous sodium sulfate and concentrated under vacuum. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (12:1). The collected fractions were combined and concentrated under vacuum. This gave 72 mg (36%) of 2-[2-[3-(5-[1-[(2S,4R)-4-hydroxy-2-([[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]carbamoyl)pyrrolidin-1-yl]-3-methyl-1-oxobutan-2-yl]-1,2-oxazol-3-yl)propoxy]ethoxy]ethyl 4-methylbenzene-1-sulfonate as a yellow oil.
工程11
50mLの丸底フラスコに、2-[2-[3-(5-[1-[(2S,4R)-4-ヒドロキシ-2-([[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]カルバモイル)ピロリジン-1-イル]-3-メチル-1-オキソブタン-2-イル]-1,2-オキサゾール-3-イル)プロポキシ]エトキシ]エチル4-メチルベンゼン-1-スルホネート(70mg、0.09mmol、1.00当量)、6-[2-(メトキシメトキシ)フェニル]-4-(1H-ピラゾール-4-イル)ピリダジン-3-アミン(27mg、0.09mmol、1.00当量)、アセトニトリル(2mL)、炭酸カリウム(38mg、0.27mmol、3.00当量)を入れた。得られた溶液を80℃で一晩撹拌した。得られた溶液を酢酸エチルで抽出し、水で洗浄した。混合物を、無水硫酸ナトリウム上で乾燥させ、真空下で濃縮した。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。収集した画分をまとめ、真空下で濃縮した。これにより、30mg(37%)の(2S,4R)-1-[2-[3-(3-[2-[2-(4-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-1H-ピラゾール-1-イル)エトキシ]エトキシ]プロピル)-1,2-オキサゾール-5-イル]-3-メチルブタノイル]-4-ヒドロキシ-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]ピロリジン-2-カルボキサミドを黄色の油として得た。 A 50 mL round-bottom flask was charged with 2-[2-[3-(5-[1-[(2S,4R)-4-hydroxy-2-([[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]carbamoyl)pyrrolidin-1-yl]-3-methyl-1-oxobutan-2-yl]-1,2-oxazol-3-yl)propoxy]ethoxy]ethyl 4-methylbenzene-1-sulfonate (70 mg, 0.09 mmol, 1.00 equiv.), 6-[2-(methoxymethoxy)phenyl]-4-(1H-pyrazol-4-yl)pyridazin-3-amine (27 mg, 0.09 mmol, 1.00 equiv.), acetonitrile (2 mL), and potassium carbonate (38 mg, 0.27 mmol, 3.00 equiv.). The resulting solution was stirred at 80°C overnight. The resulting solution was extracted with ethyl acetate and washed with water. The mixture was dried over anhydrous sodium sulfate and concentrated under vacuum. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (10:1). The collected fractions were combined and concentrated under vacuum. This gave 30 mg (37%) of (2S,4R)-1-[2-[3-(3-[2-[2-(4-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-1H-pyrazol-1-yl)ethoxy]ethoxy]propyl)-1,2-oxazol-5-yl]-3-methylbutanoyl]-4-hydroxy-N-[[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]pyrrolidine-2-carboxamide as a yellow oil.
工程12

50mLの丸底フラスコに、(2S,4R)-1-[2-[3-(3-[2-[2-(4-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-1H-ピラゾール-1-イル)エトキシ]エトキシ]プロピル)-1,2-オキサゾール-5-イル]-3-メチルブタノイル]-4-ヒドロキシ-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]ピロリジン-2-カルボキサミド(27.0mg、0.03mmol、1.00当量)、メタノール(2mL)、塩化水素(水溶液)(0.5mL)を入れた。得られた溶液を室温で一晩撹拌した。得られた溶液を水で希釈した。溶液のpH値を、炭酸ナトリウムを用いて8に調整した。得られた溶液を酢酸エチルで抽出し、水で洗浄した。混合物を、無水硫酸ナトリウム上で乾燥させ、真空下で濃縮した。以下の条件の分取HPLCによって、粗生成物を精製した:カラム:XBridge Shield RP18 OBD Column、5um、19*150mm;移動相A:水(0.05%NH3H2O)、移動相B:アセトニトリル、流速:20mL/分、勾配:8分間で34%B~47%B、220nm。これにより、7.6mg(30%)の(2S,4R)-1-[2-(3-[3-[2-(2-[4-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-1H-ピラゾール-1-イル]エトキシ)エトキシ]プロピル]-1,2-オキサゾール-5-イル)-3-メチルブタノイル]-4-ヒドロキシ-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]ピロリジン-2-カルボキサミドを白色の固体として得た。 A 50 mL round-bottom flask was charged with (2S,4R)-1-[2-[3-(3-[2-[2-(4-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-1H-pyrazol-1-yl)ethoxy]ethoxy]propyl)-1,2-oxazol-5-yl]-3-methylbutanoyl]-4-hydroxy-N-[[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]pyrrolidine-2-carboxamide (27.0 mg, 0.03 mmol, 1.00 equiv.), methanol (2 mL), and hydrogen chloride (aq) (0.5 mL). The resulting solution was stirred at room temperature overnight. The resulting solution was diluted with water. The pH value of the solution was adjusted to 8 using sodium carbonate. The resulting solution was extracted with ethyl acetate and washed with water. The mixture was dried over anhydrous sodium sulfate and concentrated under vacuum. The crude product was purified by preparative HPLC with the following conditions: Column: XBridge Shield RP18 OBD Column, 5um, 19*150mm; Mobile phase A: Water (0.05% NH3H2O), Mobile phase B: Acetonitrile, Flow rate: 20mL/min, Gradient: 34%B to 47%B in 8min, 220nm. This gave 7.6 mg (30%) of (2S,4R)-1-[2-(3-[3-[2-(2-[4-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-1H-pyrazol-1-yl]ethoxy)ethoxy]propyl]-1,2-oxazol-5-yl)-3-methylbutanoyl]-4-hydroxy-N-[[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]pyrrolidine-2-carboxamide as a white solid.
1H-NMR:(400MHz、CD3OD)δ8.89~8.83(m、1H)、8.31~8.26(m、1H)、8.08~8.03(m、2H)、7.85~7.81(m、1H)、7.46~7.21(m、5H)、6.96~6.87(m、2H)、6.15(s、1H)、4.52~4.44(m、5H)、4.41~4.31(m、3H)、3.91~3.80(m、2H)、3.72~3.65(m、1H)、3.61~3.35(m、6H)、2.62~2.51(m、2H)、2.45~2.33(m、3H)、2.22~2.15(m、1H)、2.10~2.01(m、1H)、1.84~1.72(m、2H)、1.30~1.24(m、1H)、1.03~0.97(m、3H)、0.91~0.75(m、3H)。 1H-NMR: (400MHz, CD3OD) δ 8.89-8.83 (m, 1H), 8.31-8.26 (m, 1H), 8.08-8.03 (m, 2H), 7. 85-7.81 (m, 1H), 7.46-7.21 (m, 5H), 6.96-6.87 (m, 2H), 6.15 (s, 1H), 4.52- 4.44 (m, 5H), 4.41-4.31 (m, 3H), 3.91-3.8 0 (m, 2H), 3.72-3.65 (m, 1H), 3.61-3.35 (m, 6H), 2.62-2.51 (m, 2H), 2.45- 2.33 (m, 3H), 2.22-2.15 (m, 1H), 2.10-2.01 (m, 1H), 1.84-1.72 (m, 2H), 1. 30-1.24 (m, 1H), 1.03-0.97 (m, 3H), 0.91-0.75 (m, 3H).
例示的な化合物23および例示的な化合物24の例示的な合成
25mLの丸底フラスコに、(2S,4R)-1-[2-[3-(3-[2-[2-(2-[4-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-1H-ピラゾール-1-イル]エトキシ)エトキシ]エトキシ]プロピル)-1,2-オキサゾール-5-イル]-3-メチルブタノイル]-4-ヒドロキシ-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]ピロリジン-2-カルボキサミド(87mg、0.10mmol、1.00当量)[例示的な化合物22について記載されるように調製]およびメタノール(10mL)を入れた。得られた溶液を25℃で1時間撹拌した。粗生成物を、以下の条件を用いて、キラル分取HPLCによって精製した。カラム、CHIRALPAK ID-3;移動相、MtBE(0.1%DEA):EtOH=80:20、サイズ:0.46*10cm;3um;検出器、UV-254nm。これにより、17mg(20%)の(2S,4R)-1-[(2S)-2-[3-(3-[2-[2-(2-[4-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-1H-ピラゾール-1-イル]エトキシ)エトキシ]エトキシ]プロピル)-1,2-オキサゾール-5-イル]-3-メチルブタノイル]-4-ヒドロキシ-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]ピロリジン-2-カルボキサミドを灰色がかった白色の固体[1H NMR(300MHz、CD3OD、ppm):δ8.83(s、1H)、8.33(s、1H)、8.23(s、1H)、7.99(s、1H)、7.70~7.67(d、J=9Hz、1H)、7.48~7.45(d、J=9Hz、1H)、7.37~7.32(m、4H)、7.00~6.97(m、2H)、6.19(s、1H)、4.61~4.58(m、1H)、4.50~4.38(m、5H)、3.91~3.86(m、3H)、3.60~3.51(m、8H)、3.50~3.43(m、2H)、3.40~3.34(m、2H)、2.57~2.52(m、2H)、2.46~2.43(m、4H)、2.30~2.23(m、1H)、2.10~2.04(m、1H)、1.73~1.69(m、2H)、1.07~1.05(d、J=6.6Hz、3H)、0.88~0.86(d、J=6.9Hz、3H)]として、および21mg(24%)の(2S,4R)-1-[(2R)-2-[3-(3-[2-[2-(2-[4-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-1H-ピラゾール-1-イル]エトキシ)エトキシ]エトキシ]プロピル)-1,2-オキサゾール-5-イル]-3-メチルブタノイル]-4-ヒドロキシ-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]ピロリジン-2-カルボキサミドを白色の固体[1H NMR(300MHz、CD3OD、ppm):δ8.88(s、1H)、8.37(s、1H)、8.31(s、1H)、8.04(s、1H)、7.69~7.67(d、J=8.1Hz、ppm)、7.43~7.39(m、5H)、7.03~6.99(m、2H)、6.21~6.00(m、1H)、4.53~4.48(m、3H)、4.44~4.40(m、3H)、3.92~3.89(m、3H)、3.79~3.76(m、1H)、3.62~3.54(m、7H)、3.50~3.48(m、2H)、3.42~3.40(m、2H)、2.64~2.61(m、2H)、2.46~2.44(m、4H)、2.26~2.18(m、1H)、2.13~2.04(m、1H)、1.82~1.78(m、2H)、1.05~1.02(d、J=6.6Hz、3H)、0.85~0.82(d、J=6.9Hz、3H)として得た。 A 25 mL round bottom flask was charged with (2S,4R)-1-[2-[3-(3-[2-[2-(2-[4-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-1H-pyrazol-1-yl]ethoxy)ethoxy]ethoxy]propyl)-1,2-oxazol-5-yl]-3-methylbutanoyl]-4-hydroxy-N-[[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]pyrrolidine-2-carboxamide (87 mg, 0.10 mmol, 1.00 equiv.) [prepared as described for exemplary compound 22] and methanol (10 mL). The resulting solution was stirred at 25° C. for 1 h. The crude product was purified by chiral preparative HPLC using the following conditions: Column, CHIRALPAK ID-3; mobile phase, MtBE (0.1% DEA):EtOH=80:20, size: 0.46*10 cm; 3 um; detector, UV-254 nm. This gave 17 mg (20%) of (2S,4R)-1-[(2S)-2-[3-(3-[2-[2-(2-[4-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-1H-pyrazol-1-yl]ethoxy)ethoxy]ethoxy]propyl)-1,2-oxazol-5-yl]-3-methylbutanoyl]-4-hydroxy-N-[[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]pyrrolidine-2-carboxamide as an off-white solid [1H NMR (300 MHz, CD3OD, ppm): δ 8.83 (s, 1H), 8.33 (s, 1H), 8.23 (s, 1H), 7.99 (s, 1H), 7.70-7.67 (d, J = 9 Hz, 1H), 7.48-7.45 (d, J = 9 Hz, 1H), 7.37-7.32 (m, 4H), 7.00-6.97 (m, 2H), 6.19 (s , 1H), 4.61-4.58 (m, 1H), 4.50-4.38 (m, 5H), 3.91-3.86 (m, 3H), 3.60-3.51 (m, 8H), 3.50-3.43 (m, 2H), 3.40-3.34 (m, 2H), 2.57-2.52 (m, 2H), 2.46-2.43 (m, 4H), 2.30-2.23 (m, 1H), 2 10-2.04 (m, 1H), 1.73-1.69 (m, 2H), 1.07-1.05 (d, J = 6.6 Hz, 3H), 0.88-0.86 (d, J = 6.9 Hz, 3H)] and 21 mg (24%) of (2S,4R)-1-[(2R)-2-[3-(3-[2-[2-(2-[4-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-1H-pyrazol-1-yl]ethoxy)ethoxy]ethoxy]propyl)-1,2-oxazol-5-yl]-3-methylbutanoyl]-4-hydroxy-N-[[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]pyrrolidine-2-carboxamide as a white solid [1H NMR (300 MHz, CD3OD, ppm): δ 8.88 (s, 1H), 8.37 (s, 1H), 8.31 (s, 1H), 8.04 (s, 1H), 7.69-7.67 (d, J = 8.1 Hz, ppm), 7.43-7.39 (m, 5H), 7.03-6.99 (m, 2H), 6.21-6.00 (m, 1H), 4.53-4.48 (m, 3H), 4.44-4.40 (m, 3H), 3.92-3.89 (m, 3H), 3.79- Obtained as 3.76 (m, 1H), 3.62-3.54 (m, 7H), 3.50-3.48 (m, 2H), 3.42-3.40 (m, 2H), 2.64-2.61 (m, 2H), 2.46-2.44 (m, 4H), 2.26-2.18 (m, 1H), 2.13-2.04 (m, 1H), 1.82-1.78 (m, 2H), 1.05-1.02 (d, J = 6.6 Hz, 3H), 0.85-0.82 (d, J = 6.9 Hz, 3H).
例示的な化合物25および26を、例示的な化合物22、23、および24について上に記載されるものと類似の手順を使用して調製した。 Exemplary compounds 25 and 26 were prepared using procedures similar to those described above for exemplary compounds 22, 23, and 24.
例示的な化合物27の例示的な合成
工程1
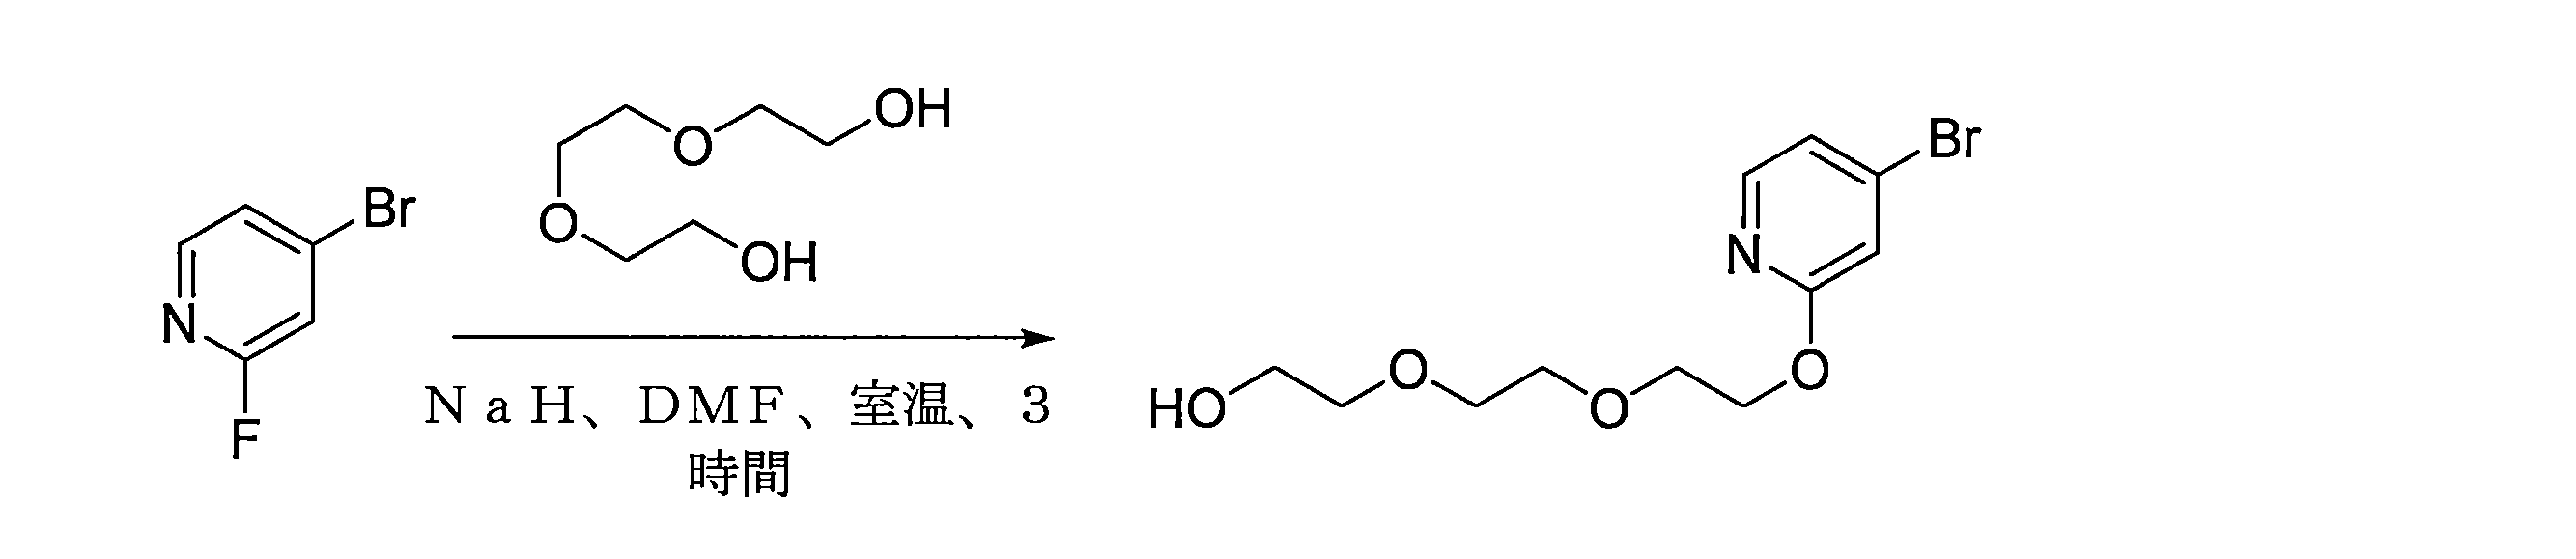
50mLの丸底フラスコに、2-[2-(2-ヒドロキシエトキシ)エトキシ]エタン-1-オール(257mg、1.71mmol、3.00当量)およびN,N-ジメチルホルムアミド(5mL)を入れた。これに続いて、水素化ナトリウム(34mg、1.42mmol、1.50当量)を0℃で10分間添加した。これに、4-ブロモ-2-フルオロピリジン(100mg、0.57mmol、1.00当量)を添加した。得られた溶液を室温で3時間撹拌した。その後、5mLの水/氷を添加することによって反応を停止させた。得られた溶液を酢酸エチルで抽出し、有機層を合わせた。得られた混合物を、飽和塩化ナトリウムで洗浄した。混合物を、無水硫酸ナトリウムで乾燥させた。残渣を、酢酸エチル/石油エーテル(1:1)とともにシリカゲルカラムに充填させた。収集した画分をまとめ、真空下で濃縮した。これにより、90mg(52%)の2-(2-[2-[(4-ブロモピリジン-2-イル)オキシ]エトキシ]エトキシ)エタン-1-オールを淡い黄色の液体として得た。 A 50 mL round bottom flask was charged with 2-[2-(2-hydroxyethoxy)ethoxy]ethan-1-ol (257 mg, 1.71 mmol, 3.00 equiv.) and N,N-dimethylformamide (5 mL). This was followed by the addition of sodium hydride (34 mg, 1.42 mmol, 1.50 equiv.) at 0°C for 10 min. To this was added 4-bromo-2-fluoropyridine (100 mg, 0.57 mmol, 1.00 equiv.). The resulting solution was stirred at room temperature for 3 h. The reaction was then quenched by adding 5 mL of water/ice. The resulting solution was extracted with ethyl acetate and the organic layers were combined. The resulting mixture was washed with saturated sodium chloride. The mixture was dried over anhydrous sodium sulfate. The residue was loaded onto a silica gel column with ethyl acetate/petroleum ether (1:1). The collected fractions were combined and concentrated under vacuum. This yielded 90 mg (52%) of 2-(2-[2-[(4-bromopyridin-2-yl)oxy]ethoxy]ethoxy)ethan-1-ol as a pale yellow liquid.
工程2

窒素不活性雰囲気下でパージし、維持した50mLの丸底フラスコに、2-(2-[2-[(4-ブロモピリジン-2-イル)オキシ]エトキシ]エトキシ)エタン-1-オール(500mg、1.63mmol、1.00当量)、tert-ブチル3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレート(347mg、1.64mmol、1.00当量)、Cs2CO3(1599mg、4.91mmol、3.00当量)、トルエン(8mL)、Ruphos(69mg、0.05当量)を入れた。得られた溶液を、油浴中で、100℃で5時間撹拌した。次いで、反応を、10mLの水の添加により停止させた。得られた溶液をジクロロメタン/MeOHで抽出し、有機層を合わせた。得られた混合物を、飽和塩化ナトリウムで洗浄した。混合物を、無水硫酸ナトリウムで乾燥させた。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。収集した画分をまとめ、真空下で濃縮した。これにより、419mg(59%)のtert-ブチル8-(2-[2-[2-(2-ヒドロキシエトキシ)エトキシ]エトキシ]ピリジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレートを黄色の油として得た。 A 50 mL round bottom flask purged and maintained under a nitrogen inert atmosphere was charged with 2-(2-[2-[(4-bromopyridin-2-yl)oxy]ethoxy]ethoxy)ethan-1-ol (500 mg, 1.63 mmol, 1.00 equiv.), tert-butyl 3,8-diazabicyclo[3.2.1]octane-3-carboxylate (347 mg, 1.64 mmol, 1.00 equiv.), Cs2CO3 (1599 mg, 4.91 mmol, 3.00 equiv.), toluene (8 mL), and Ruphos (69 mg, 0.05 equiv.). The resulting solution was stirred in an oil bath at 100°C for 5 hours. The reaction was then quenched by the addition of 10 mL of water. The resulting solution was extracted with dichloromethane/MeOH and the organic layers were combined. The resulting mixture was washed with saturated sodium chloride. The mixture was dried over anhydrous sodium sulfate. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (10:1). The collected fractions were combined and concentrated under vacuum. This gave 419 mg (59%) of tert-butyl 8-(2-[2-[2-(2-hydroxyethoxy)ethoxy]ethoxy]pyridin-4-yl)-3,8-diazabicyclo[3.2.1]octane-3-carboxylate as a yellow oil.
工程3

50mLの丸底フラスコに、tert-ブチル8-(2-[2-[2-(2-ヒドロキシエトキシ)エトキシ]エトキシ]ピリジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレート(419mg、0.96mmol、1.00当量)およびメタノール中の1M HCl(8mL)を入れた。得られた溶液を室温で2時間撹拌した。得られた混合物を真空下で濃縮した。これにより、319mg(99%)の2-(2-[2-[(4-[3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)オキシ]エトキシ]エトキシ)エタン-1-オールを黄色の油として得た。 A 50 mL round bottom flask was charged with tert-butyl 8-(2-[2-[2-(2-hydroxyethoxy)ethoxy]ethoxy]pyridin-4-yl)-3,8-diazabicyclo[3.2.1]octane-3-carboxylate (419 mg, 0.96 mmol, 1.00 equiv.) and 1 M HCl in methanol (8 mL). The resulting solution was stirred at room temperature for 2 h. The resulting mixture was concentrated under vacuum. This afforded 319 mg (99%) of 2-(2-[2-[(4-[3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl)oxy]ethoxy]ethoxy)ethan-1-ol as a yellow oil.
工程4

10mLのマイクロ波チューブに、2-(2-[2-[(4-[3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)オキシ]エトキシ]エトキシ)エタン-1-オール(319mg、0.95mmol、1.00当量)、4-ブロモ-6-クロロピリダジン-3-アミン(780mg、3.74mmol、4.00当量)、DMSO(10mL)、DIEA(2mL)を入れた。最終反応混合物を、マイクロ波放射で、130℃で3時間照射した。次いで、反応物を、10mLの水を添加することによってクエンチした。得られた溶液を酢酸エチルで抽出し、有機層を合わせた。得られた混合物を、飽和塩化ナトリウムで洗浄した。混合物を、無水硫酸ナトリウムで乾燥させた。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。収集した画分をまとめ、真空下で濃縮した。これにより、260mg(59%)の2-[2-[2-([4-[3-(3-アミノ-6-クロロピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル]オキシ)エトキシ]エトキシ]エタン-1-オールを黄色の固体として得た。 A 10 mL microwave tube was charged with 2-(2-[2-[(4-[3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl)oxy]ethoxy]ethoxy)ethan-1-ol (319 mg, 0.95 mmol, 1.00 equiv.), 4-bromo-6-chloropyridazin-3-amine (780 mg, 3.74 mmol, 4.00 equiv.), DMSO (10 mL), and DIEA (2 mL). The final reaction mixture was irradiated with microwave radiation at 130° C. for 3 hours. The reaction was then quenched by adding 10 mL of water. The resulting solution was extracted with ethyl acetate and the organic layers were combined. The resulting mixture was washed with saturated sodium chloride. The mixture was dried over anhydrous sodium sulfate. The residue was applied onto a silica gel column eluting with dichloromethane/methanol (10:1). The collected fractions were combined and concentrated under vacuum. This gave 260 mg (59%) of 2-[2-[2-([4-[3-(3-amino-6-chloropyridazin-4-yl)-3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl]oxy)ethoxy]ethoxy]ethan-1-ol as a yellow solid.
工程5
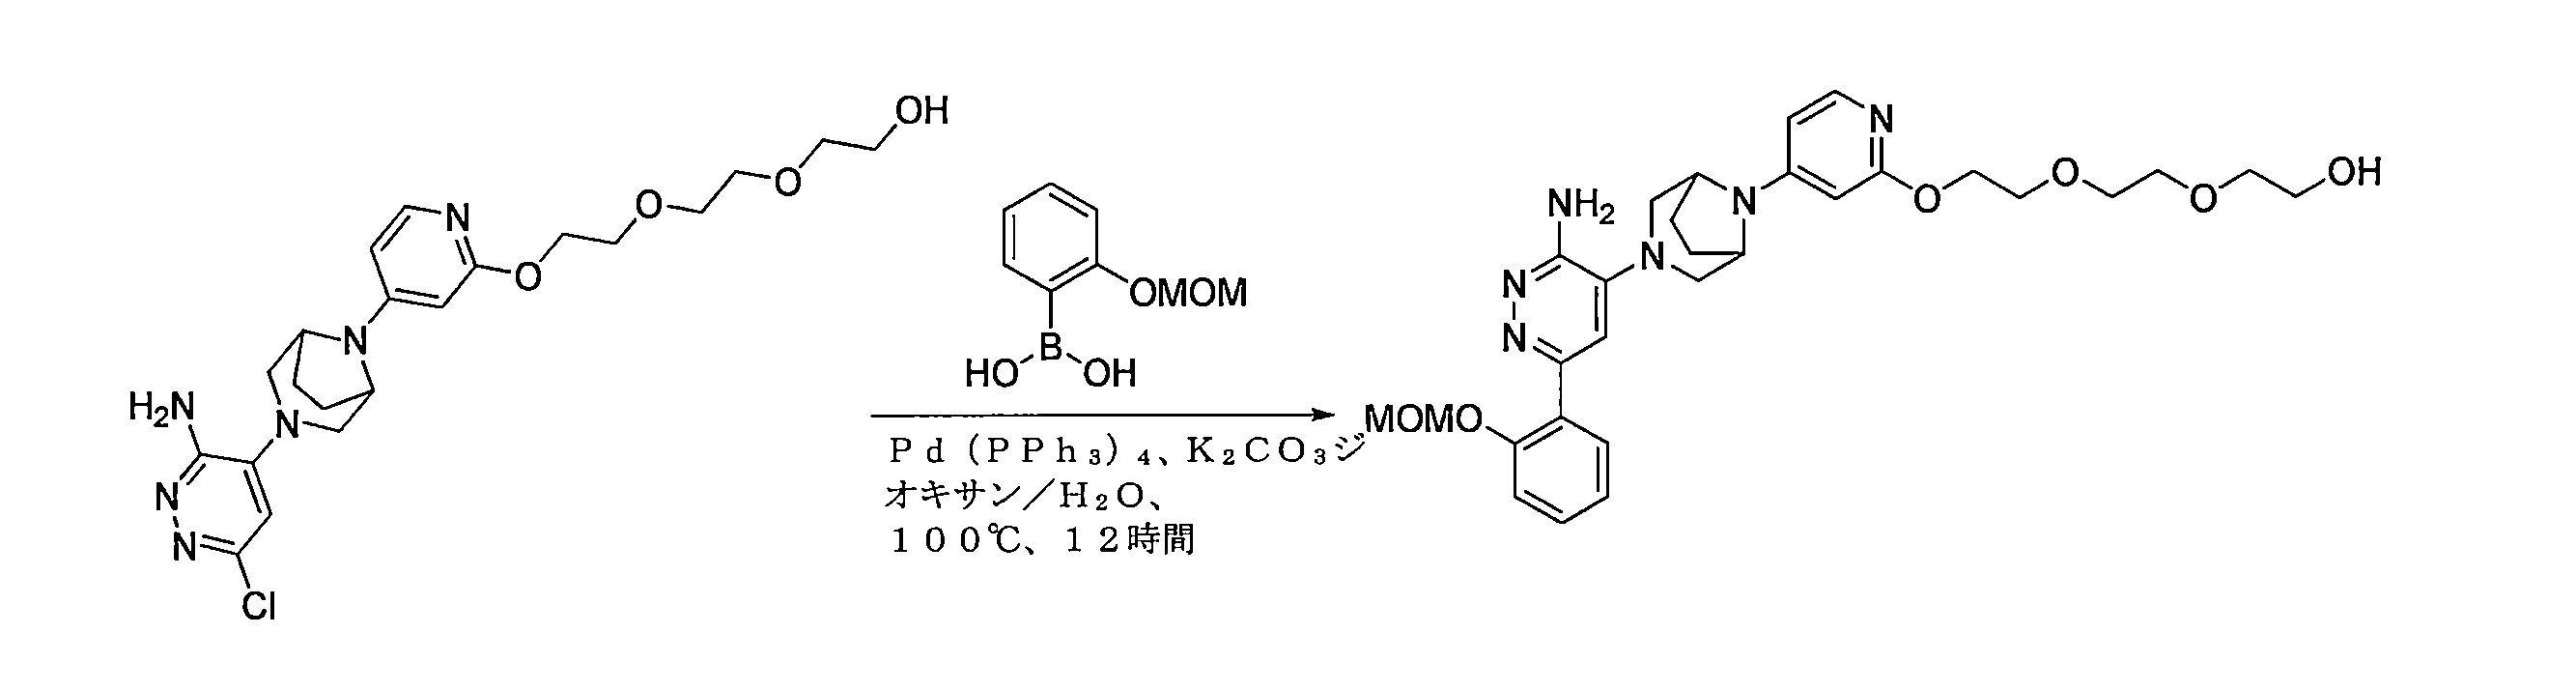
窒素不活性雰囲気下でパージし、維持した10mLのマイクロ波チューブに、2-[2-[2-([4-[3-(3-アミノ-6-クロロピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル]オキシ)エトキシ]エトキシ]エタン-1-オール(295mg、0.63mmol、1.00当量)、[2-(メトキシメトキシ)フェニル]ボロン酸(223mg、1.23mmol、2.00当量)、炭酸カリウム(254mg、1.84mmol、3.00当量)、ジオキサン(4mL)、水(1mL)、Pd(PPh3)4(70mg、0.06mmol、0.10当量)を入れた。得られた溶液を100℃で一晩撹拌した。次いで、反応物を、水を添加することによってクエンチした。得られた溶液をジクロロメタン/MeOHで抽出し、有機層を合わせた。得られた混合物を、飽和塩化ナトリウムで洗浄した。混合物を、無水硫酸ナトリウムで乾燥させた。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。収集した画分をまとめ、真空下で濃縮した。これにより、221mg(61%)の2-[2-(2-[[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]オキシ]エトキシ)エトキシ]エタン-1-オールを黄色の固体として得た。 A 10 mL microwave tube purged and maintained under a nitrogen inert atmosphere was charged with 2-[2-[2-([4-[3-(3-amino-6-chloropyridazin-4-yl)-3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl]oxy)ethoxy]ethoxy]ethan-1-ol (295 mg, 0.63 mmol, 1.00 equiv.), [2-(methoxymethoxy)phenyl]boronic acid (223 mg, 1.23 mmol, 2.00 equiv.), potassium carbonate (254 mg, 1.84 mmol, 3.00 equiv.), dioxane (4 mL), water (1 mL), and Pd(PPh3)4 (70 mg, 0.06 mmol, 0.10 equiv.). The resulting solution was stirred at 100° C. overnight. The reaction was then quenched by adding water. The resulting solution was extracted with dichloromethane/MeOH and the organic layers were combined. The resulting mixture was washed with saturated sodium chloride. The mixture was dried over anhydrous sodium sulfate. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (10:1). The collected fractions were combined and concentrated under vacuum. This gave 221 mg (61%) of 2-[2-(2-[[4-(3-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl]oxy]ethoxy)ethoxy]ethan-1-ol as a yellow solid.
工程6

50mLの丸底フラスコに、2-[2-(2-[[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]オキシ]エトキシ)エトキシ]エタン-1-オール(100mg、0.18mmol、1.00当量)、TsCl(50mg、0.26mmol、1.50当量)、ジクロロメタン(5mL)、トリエチルアミン(0.3mL)、4-ジメチルアミノピリジン(2mg、0.02mmol、0.10当量)を入れた。得られた溶液を室温で2時間撹拌した。次いで、反応を、水の添加により停止させた。得られた溶液を酢酸エチルで抽出し、有機層を合わせた。得られた混合物を、飽和塩化ナトリウムで洗浄した。混合物を、無水硫酸ナトリウムで乾燥させた。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。収集した画分をまとめ、真空下で濃縮した。これにより、97mg(76%)の2-[2-(2-[[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]オキシ]エトキシ)エトキシ]エチル4-メチルベンゼン-1-スルホネートを黄色の固体として得た。 A 50 mL round bottom flask was charged with 2-[2-(2-[[4-(3-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl]oxy]ethoxy)ethoxy]ethan-1-ol (100 mg, 0.18 mmol, 1.00 equiv.), TsCl (50 mg, 0.26 mmol, 1.50 equiv.), dichloromethane (5 mL), triethylamine (0.3 mL), and 4-dimethylaminopyridine (2 mg, 0.02 mmol, 0.10 equiv.). The resulting solution was stirred at room temperature for 2 hours. The reaction was then quenched by the addition of water. The resulting solution was extracted with ethyl acetate and the organic layers were combined. The resulting mixture was washed with saturated sodium chloride. The mixture was dried over anhydrous sodium sulfate. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (10:1). The collected fractions were combined and concentrated under vacuum. This gave 97 mg (76%) of 2-[2-(2-[[4-(3-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl]oxy]ethoxy)ethoxy]ethyl 4-methylbenzene-1-sulfonate as a yellow solid.
工程7

50mLの丸底フラスコに、2-[2-(2-[[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]オキシ]エトキシ)エトキシ]エチル4-メチルベンゼン-1-スルホネート(87mg、0.12mmol、1.00当量)、N,N-ジメチルホルムアミド(54mg、0.75mmol、1.00当量)、(2S,4R)-4-ヒドロキシ-1-[2-(3-ヒドロキシ-1,2-オキサゾール-5-イル)-3-メチルブタノイル]-N-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチルピロリジン-2-カルボキサミド(79mg、0.16mmol、2.00当量)、Cs2CO3(5g、15.35mmol、127.15当量)を入れた。得られた溶液を室温で4時間撹拌した。得られた溶液をジクロロメタン/MeOHで抽出し、有機層を合わせた。得られた混合物を、飽和塩化ナトリウムで洗浄した。混合物を、無水硫酸ナトリウムで乾燥させた。残渣を、ジクロロメタン/メタノール(5:1)で溶出するシリカゲルカラム上に適用した。収集した画分をまとめ、真空下で濃縮した。これにより、124mg(99%)の(2S,4R)-1-[2-(3-[2-[2-(2-[[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]オキシ]エトキシ)エトキシ]エトキシ]-1,2-オキサゾール-5-イル)-3-メチルブタノイル]-4-ヒドロキシ-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]ピロリジン-2-カルボキサミドを黄色の固体として得た。 In a 50 mL round-bottom flask, add 2-[2-(2-[[4-(3-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl]oxy]ethoxy)ethoxy]ethyl 4-methylbenzene-1-sulfonate (87 mg, 0.12 mmol, 1.00 equiv.), N,N-dimethylformamide (54 mg, 0.75 mmol, 1.00 equiv), (2S,4R)-4-hydroxy-1-[2-(3-hydroxy-1,2-oxazol-5-yl)-3-methylbutanoyl]-N-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methylpyrrolidine-2-carboxamide (79 mg, 0.16 mmol, 2.00 equiv), and Cs2CO3 (5 g, 15.35 mmol, 127.15 equiv). The resulting solution was stirred at room temperature for 4 hours. The resulting solution was extracted with dichloromethane/MeOH and the organic layers were combined. The resulting mixture was washed with saturated sodium chloride. The mixture was dried over anhydrous sodium sulfate. The residue was applied onto a silica gel column eluting with dichloromethane/methanol (5:1). The collected fractions were combined and concentrated under vacuum. This gave 124 mg (99%) of (2S,4R)-1-[2-(3-[2-[2-(2-[[4-(3-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl]oxy]ethoxy)ethoxy]ethoxy]-1,2-oxazol-5-yl)-3-methylbutanoyl]-4-hydroxy-N-[[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]pyrrolidine-2-carboxamide as a yellow solid.
工程8

50mLの丸底フラスコに、(2S,4R)-1-[2-(3-[2-[2-(2-[[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]オキシ]エトキシ)エトキシ]エトキシ]-1,2-オキサゾール-5-イル)-3-メチルブタノイル]-4-ヒドロキシ-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]ピロリジン-2-カルボキサミド(131mg、0.13mmol、1.00当量)、メタノール(5mL)、およびMeOH中の1M HCl(1.5mL)を入れた。得られた溶液を、室温で7時間撹拌した。得られた溶液を5mLのH2Oで希釈した。溶液のpH値を、Na2CO3水溶液(2M)で7に調整した。得られた溶液をジクロロメタンで抽出し、有機層を合わせた。得られた混合物を、飽和塩化ナトリウムで洗浄した。以下の条件の分取HPLCによって、粗生成物を精製した:カラム、XBridge Shield RP18 OBDカラム、5um、19*150mm;移動相、水(0.05%NH3H2O)およびアセトニトリル(39.0%アセトニトリル、9分間で最大50.0%);検出器、UV220nm。これにより、60mg(48%)の(2S,4R)-1-(2-[3-[2-(2-[2-[(4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)オキシ]エトキシ]エトキシ)エトキシ]-1,2-オキサゾール-5-イル]-3-メチルブタノイル)-4-ヒドロキシ-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]ピロリジン-2-カルボキサミドを黄色の固体として得、これを、以下の条件を用いて、キラル分取HPLCによってさらに精製した。カラム、Chiralpak ID-2、2*25cm、5um;移動相、(0.1%DEA)およびエタノール(30分間で30%エタノールを保持);検出器、UV254/220nm。これにより、13.9mg(23%)の(2S,4R)-1-[(2S)-2-[3-[2-(2-[2-[(4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)オキシ]エトキシ]エトキシ)エトキシ]-1,2-オキサゾール-5-イル]-3-メチルブタノイル]-4-ヒドロキシ-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]ピロリジン-2-カルボキサミドを白色の固体[1H NMR(400MHz、CD3OD):δ8.85(m、1H)、7.78~7.72(m、2H)、7.47~7.45(m、2H)、7.40~7.34(m、3H)、7.23~7.22(m、1H)、6.92~6.88(m、2H)、6.55~6.53(m、1H)、6.23(s、1H)、5.97(s、1H)、4.64~4.59(m、2H)、4.50(s、3H)、4.40(s、2H)、4.33~4.24(m、4H)、3.81~3.79(m、2H)、3.75~3.65(m、8H)、3.29(m、1H)、3.10~3.07(m、2H)、2.48~2.46(m、3H)、2.23~2.20(m,1H)、2.13~2.06(m、6H)、1.05(s、1H)、0.90~0.89(d、J=6.8Hz、3H)、0.78~0.76(d、J=8Hz、3H)]として、および17.9mg(29%)の(2S,4R)-1-[(2R)-2-[3-[2-(2-[2-[(4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)オキシ]エトキシ]エトキシ)エトキシ]-1,2-オキサゾール-5-イル]-3-メチルブタノイル]-4-ヒドロキシ-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]ピロリジン-2-カルボキサミドを白色の固体[1H NMR(400MHz、CD3OD):δ8.88(m、1H)、7.79~7.75(m、2H)、7.49~7.39(m、5H)、7.24(m、1H)、6.92~6.90(m、2H)、6.56~6.54(m、1H)、6.25~6.24(M、1H)、6.00(s、1H)、4.53~4.29(m、10H)、3.88~3.81(m、5H)、3.70~3.63(m、6H)、3.33(m、2H)、3.14~3.12(m、2H)、2.48~2.44(m、3H)、2.42~2.29(m、1H)、2.25~2.12(m、6H)、1.04~1.02(d、J=6.4Hz、3H)、0.88~0.86(d、J=6.4Hz、3H)]として得た。 A 50 mL round bottom flask was charged with (2S,4R)-1-[2-(3-[2-[2-(2-[[4-(3-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl]oxy]ethoxy)ethoxy]ethoxy]-1,2-oxazol-5-yl)-3-methylbutanoyl]-4-hydroxy-N-[[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]pyrrolidine-2-carboxamide (131 mg, 0.13 mmol, 1.00 equiv.), methanol (5 mL), and 1 M HCl in MeOH (1.5 mL). The resulting solution was stirred at room temperature for 7 h. The resulting solution was diluted with 5 mL of H2O. The pH value of the solution was adjusted to 7 with aqueous Na2CO3 (2M). The resulting solution was extracted with dichloromethane and the organic layers were combined. The resulting mixture was washed with saturated sodium chloride. The crude product was purified by preparative HPLC with the following conditions: column, XBridge Shield RP18 OBD column, 5um, 19*150mm; mobile phase, water (0.05% NH3H2O) and acetonitrile (39.0% acetonitrile, max. 50.0% in 9 min); detector, UV 220nm. This gave 60 mg (48%) of (2S,4R)-1-(2-[3-[2-(2-[2-[(4-[3-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl)oxy]ethoxy]ethoxy)ethoxy]-1,2-oxazol-5-yl]-3-methylbutanoyl)-4-hydroxy-N-[[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]pyrrolidine-2-carboxamide as a yellow solid which was further purified by chiral preparative HPLC using the following conditions: Column, Chiralpak ID-2, 2*25 cm, 5 um; mobile phase, (0.1% DEA) and ethanol (30% ethanol hold for 30 min); detector, UV 254/220 nm. This gave 13.9 mg (23%) of (2S,4R)-1-[(2S)-2-[3-[2-(2-[2-[(4-[3-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl)oxy]ethoxy]ethoxy)ethoxy]-1,2-oxazol-5-yl]-3-methylbutanoyl]-4-hydroxy-N-[[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]pyrrolidine-2-carboxamide as a white solid [1H NMR (400MHz, CD3OD): δ 8.85 (m, 1H), 7.78-7.72 (m, 2H), 7.47-7.45 (m, 2H), 7.40-7.34 (m, 3H), 7.23-7.22 (m, 1H), 6.92-6.88 (m, 2H), 6.55-6.53 (m, 1H), 6.23 (s, 1H), 5.97 (s, 1H), 4.64- 4.59 (m, 2H), 4.50 (s, 3H), 4.40 (s, 2H), 4.33-4.24 (m, 4H), 3.81-3.79 (m, 2H), 3.75-3.65 (m, 8H), 3.29 (m, 1H), 3.10-3.07 (m, 2H), 2.48-2.46 (m, 3H), 2.23-2.20 (m, 1H), 2.13-2.06 (m, 6H), 3H), 1.05 (s, 1H), 0.90-0.89 (d, J = 6.8 Hz, 3H), 0.78-0.76 (d, J = 8 Hz, 3H)], and 17.9 mg (29%) of (2S,4R)-1-[(2R)-2-[3-[2-(2-[2-[(4-[3-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl)oxy]ethoxy]ethoxy)ethoxy]-1,2-oxazol-5-yl]-3-methylbutanoyl]-4-hydroxy-N-[[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]pyrrolidine-2-carboxamide as a white solid [1H NMR (400MHz, CD3OD): δ 8.88 (m, 1H), 7.79-7.75 (m, 2H), 7.49-7.39 (m, 5H), 7.24 (m, 1H), 6.92-6.90 (m, 2H), 6.56-6.54 (m, 1H), 6.25-6.24 (M, 1H), 6.00 (s, 1H), 4.53-4.29 (m, 10H), 3.88-3 . 81 (m, 5H), 3.70-3.63 (m, 6H), 3.33 (m, 2H), 3.14-3.12 (m, 2H), 2.48-2.44 (m, 3H), 2.42-2.29 (m, 1H), 2.25-2.12 (m, 6H), 1.04-1.02 (d, J = 6.4 Hz, 3H), 0.88-0.86 (d, J = 6.4 Hz, 3H).
例示的な化合物29および30を、例示的な化合物27および28について記載されるものと類似の手順を使用して調製した。 Exemplary compounds 29 and 30 were prepared using procedures similar to those described for exemplary compounds 27 and 28.
例示的な化合物31の例示的な合成
工程1

50mLの丸底フラスコに、2-(オキサン-2-イルオキシ)エタン-1-オール(4.5g、30.78mmol、1.00当量)、テトラヒドロフラン(10mL)、テトラヒドロフラン(60mL)中のt-BuOK(3.6g、32.08mmol、2.00当量)の溶液を入れた。続いて、3-ブロモプロプ-1-イン(2.61mL、1.00当量)を0℃で撹拌しながら滴下で添加した。得られた溶液を室温で一晩撹拌した。次いで、反応物を、水/氷を添加することによってクエンチした。得られた溶液を酢酸エチルで抽出し、有機層を合わせた。得られた混合物を、飽和塩化ナトリウムで洗浄した。混合物を、無水硫酸ナトリウムで乾燥させた。残渣を、酢酸エチル/石油エーテル(1:5)で溶出するシリカゲルカラム上に適用した。収集した画分をまとめ、真空下で濃縮した。これにより、2.8g(49%)の2-[2-(プロパ-2-イン-1-イルオキシ)エトキシ]オキサンを黄色の油として得た。 A 50 mL round bottom flask was charged with 2-(oxan-2-yloxy)ethan-1-ol (4.5 g, 30.78 mmol, 1.00 equiv.), tetrahydrofuran (10 mL), and a solution of t-BuOK (3.6 g, 32.08 mmol, 2.00 equiv.) in tetrahydrofuran (60 mL). Subsequently, 3-bromoprop-1-yne (2.61 mL, 1.00 equiv.) was added dropwise with stirring at 0°C. The resulting solution was stirred at room temperature overnight. The reaction was then quenched by adding water/ice. The resulting solution was extracted with ethyl acetate and the organic layers were combined. The resulting mixture was washed with saturated sodium chloride. The mixture was dried over anhydrous sodium sulfate. The residue was applied onto a silica gel column eluting with ethyl acetate/petroleum ether (1:5). The collected fractions were combined and concentrated under vacuum. This yielded 2.8 g (49%) of 2-[2-(prop-2-yn-1-yloxy)ethoxy]oxane as a yellow oil.
工程2

窒素不活性雰囲気下でパージし、維持した10mLのマイクロ波チューブに、2-[2-(プロパ-2-イン-1-イルオキシ)エトキシ]オキサン(2.8g、15.20mmol、1.00当量)、ZrCp2HCl(390mg、0.10当量)、トリエチルアミン(153mg、1.52mmol、0.10当量)、ピナコールボラン(2.5mL)を入れた。結果として生じる溶液を油浴中、68℃で一晩撹拌した。次いで、反応物を、NH4Clを添加することによってクエンチした。反応混合物を、水/氷浴で0℃に冷却した。得られた溶液を酢酸エチルで抽出し、有機層を合わせた。得られた混合物を、飽和塩化ナトリウムで洗浄した。混合物を、無水硫酸ナトリウムで乾燥させた。残渣を、酢酸エチル/石油エーテル(1:5)で溶出するシリカゲルカラム上に適用した。収集した画分をまとめ、真空下で濃縮した。これにより、3.01g(63%)の4,4,5,5-テトラメチル-2-[(1E)-3-[2-(オキサン-2-イルオキシ)エトキシ]プロパ-1-エン-1-イル]-1,3,2-ジオキサボロランを黄色の液体として得た。 A 10 mL microwave tube purged and maintained under nitrogen inert atmosphere was charged with 2-[2-(prop-2-yn-1-yloxy)ethoxy]oxane (2.8 g, 15.20 mmol, 1.00 equiv.), ZrCp2HCl (390 mg, 0.10 equiv.), triethylamine (153 mg, 1.52 mmol, 0.10 equiv.), and pinacolborane (2.5 mL). The resulting solution was stirred overnight at 68°C in an oil bath. The reaction was then quenched by adding NH4Cl. The reaction mixture was cooled to 0°C in a water/ice bath. The resulting solution was extracted with ethyl acetate and the organic layers were combined. The resulting mixture was washed with saturated sodium chloride. The mixture was dried over anhydrous sodium sulfate. The residue was applied onto a silica gel column eluting with ethyl acetate/petroleum ether (1:5). The collected fractions were combined and concentrated under vacuum. This yielded 3.01 g (63%) of 4,4,5,5-tetramethyl-2-[(1E)-3-[2-(oxan-2-yloxy)ethoxy]prop-1-en-1-yl]-1,3,2-dioxaborolane as a yellow liquid.
工程3

窒素不活性雰囲気下でパージし、維持した10mLのマイクロ波チューブに、4,4,5,5-テトラメチル-2-[(1E)-3-[2-(オキサン-2-イルオキシ)エトキシ]プロパ-1-エン-1-イル]-1,3,2-ジオキサボロラン(1g、3.20mmol、1.00当量)、Pd(PPh3)4(800mg、0.69mmol、1.00当量)、炭酸カリウム(8g、57.88mmol、18.07当量)、ジオキサン(2g)、tert-ブチル8-(2-ブロモピリジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレート(902mg、2.45mmol、3.00当量)、水(252mg、0.10当量)を入れた。得られた溶液を、油浴中で、100℃で一晩撹拌した。次いで、反応を、水の添加により停止させた。得られた溶液を酢酸エチルで抽出し、有機層を合わせた。得られた混合物を、飽和塩化ナトリウムで洗浄した。混合物を、無水硫酸ナトリウムで乾燥させた。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。収集した画分をまとめ、真空下で濃縮した。これにより、639mg(42%)のtert-ブチル8-[2-[(1E)-3-[2-(オキサン-2-イルオキシ)エトキシ]プロパ-1-エン-1-イル]ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレートを黄色の油として得た。 A 10 mL microwave tube purged and maintained under nitrogen inert atmosphere was charged with 4,4,5,5-tetramethyl-2-[(1E)-3-[2-(oxan-2-yloxy)ethoxy]prop-1-en-1-yl]-1,3,2-dioxaborolane (1 g, 3.20 mmol, 1.00 equiv.), Pd(PPh3)4 (800 mg, 0.69 mmol, 1.00 equiv.), potassium carbonate (8 g, 57.88 mmol, 18.07 equiv.), dioxane (2 g), tert-butyl 8-(2-bromopyridin-4-yl)-3,8-diazabicyclo[3.2.1]octane-3-carboxylate (902 mg, 2.45 mmol, 3.00 equiv.), and water (252 mg, 0.10 equiv.). The resulting solution was stirred overnight at 100 °C in an oil bath. The reaction was then quenched by the addition of water. The resulting solution was extracted with ethyl acetate and the organic layers were combined. The resulting mixture was washed with saturated sodium chloride. The mixture was dried over anhydrous sodium sulfate. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (10:1). The collected fractions were combined and concentrated under vacuum. This gave 639 mg (42%) of tert-butyl 8-[2-[(1E)-3-[2-(oxan-2-yloxy)ethoxy]prop-1-en-1-yl]pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octane-3-carboxylate as a yellow oil.
工程4

50mLの丸底フラスコに、tert-ブチル8-[2-[(1E)-3-[2-(オキサン-2-イルオキシ)エトキシ]プロパ-1-エン-1-イル]ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレート(639mg、1.35mmol、1.00当量)、炭素上のパラジウム(200mg)、およびメタノール(15mL)を入れ、混合物を、水素雰囲気下で、室温で一晩撹拌した。固形分を濾過し、濾液を濃縮した。これにより、639mg(100%)のtert-ブチル8-(2-[3-[2-(オキサン-2-イルオキシ)エトキシ]プロピル]ピリジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレートを黄色の油として得た。 A 50 mL round bottom flask was charged with tert-butyl 8-[2-[(1E)-3-[2-(oxan-2-yloxy)ethoxy]prop-1-en-1-yl]pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octane-3-carboxylate (639 mg, 1.35 mmol, 1.00 equiv.), palladium on carbon (200 mg), and methanol (15 mL), and the mixture was stirred overnight at room temperature under a hydrogen atmosphere. The solids were filtered and the filtrate was concentrated. This gave 639 mg (100%) of tert-butyl 8-(2-[3-[2-(oxan-2-yloxy)ethoxy]propyl]pyridin-4-yl)-3,8-diazabicyclo[3.2.1]octane-3-carboxylate as a yellow oil.
工程5
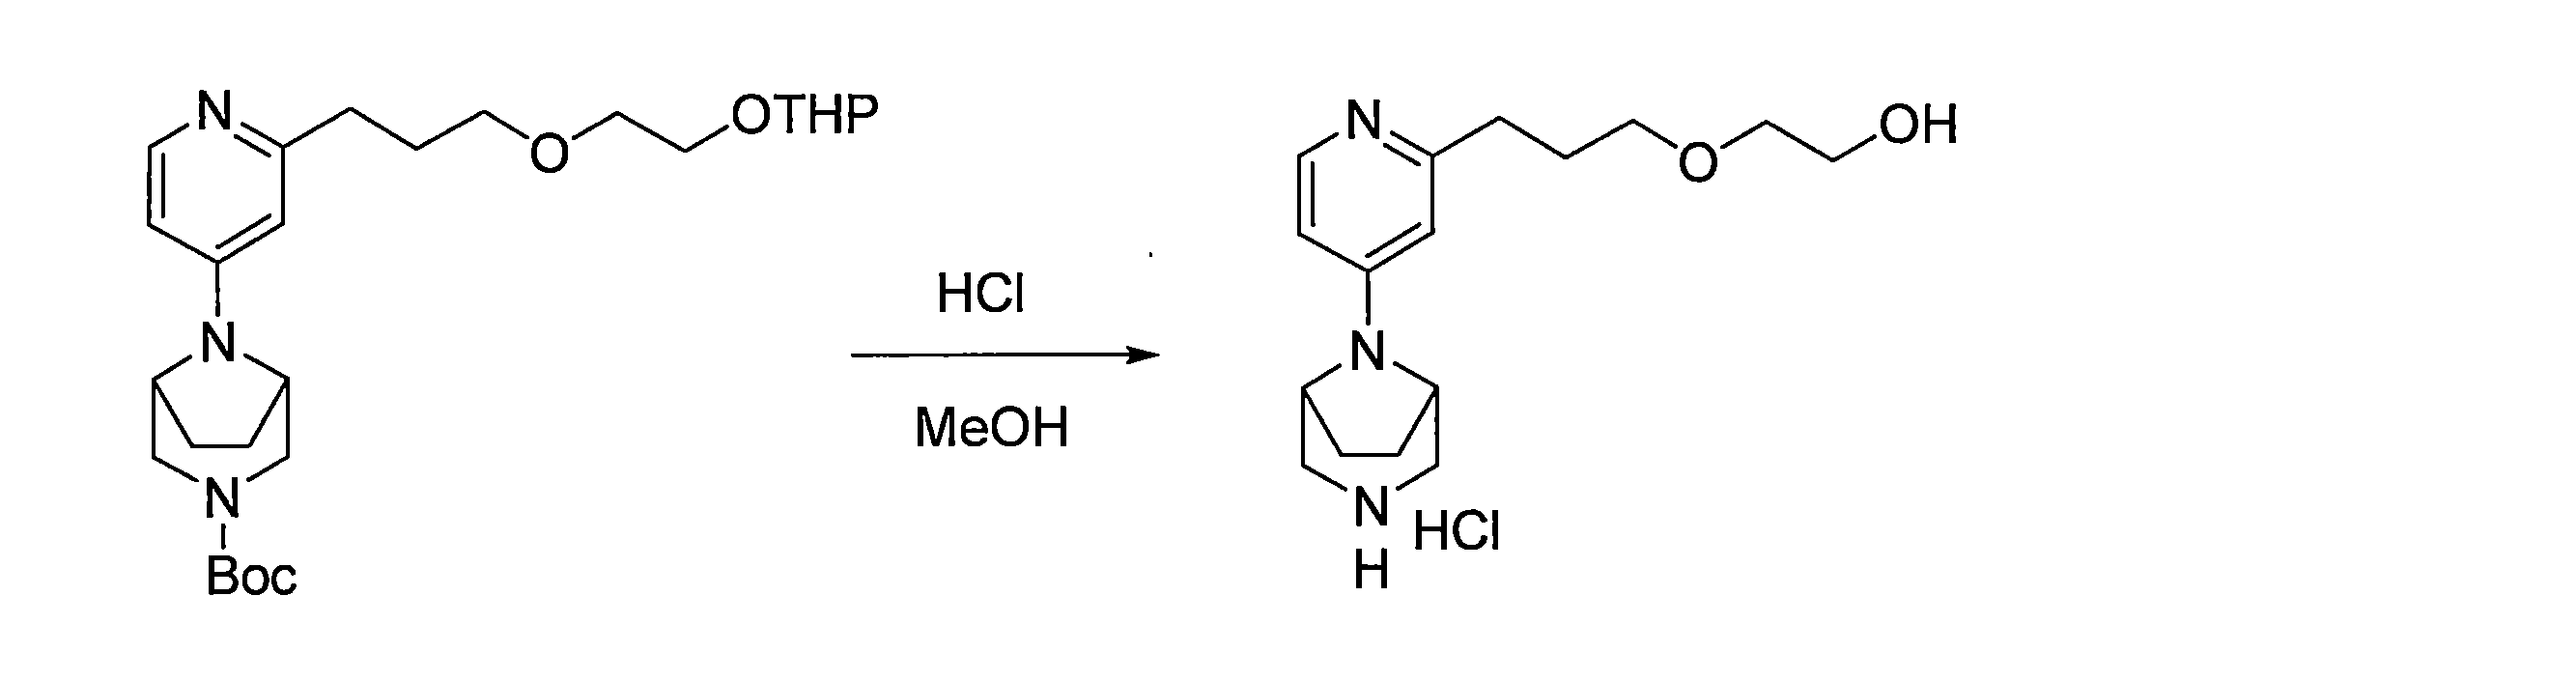
50mLの丸底フラスコに、tert-ブチル8-(2-[3-[2-(オキサン-2-イルオキシ)エトキシ]プロピル]ピリジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレート(639mg、1.34mmol、1.00当量)、メタノール(15mL)を入れ、塩化水素ガスを、溶液によって気泡化した。得られた溶液を室温で3時間撹拌した。得られた混合物を真空下で濃縮した。これにより、387mg(100%)の2-[3-(4-[3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)プロポキシ]エタン-1-オールを黄色の油として得た。 A 50 mL round bottom flask was charged with tert-butyl 8-(2-[3-[2-(oxan-2-yloxy)ethoxy]propyl]pyridin-4-yl)-3,8-diazabicyclo[3.2.1]octane-3-carboxylate (639 mg, 1.34 mmol, 1.00 equiv.) and methanol (15 mL) and hydrogen chloride gas was bubbled through the solution. The resulting solution was stirred at room temperature for 3 hours. The resulting mixture was concentrated under vacuum. This afforded 387 mg (100%) of 2-[3-(4-[3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl)propoxy]ethan-1-ol as a yellow oil.
工程6

10mLのマイクロ波チューブに、2-[3-(4-[3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)プロポキシ]エタン-1-オール(387mg、1.85mmol、1.00当量)、4-ブロモ-6-クロロピリダジン-3-アミン(1.54g、7.39mmol、4.00当量)、DMSO(8mL)、DIEA(1.53mL、5.00当量)を入れた。最終反応混合物を、マイクロ波放射で、130℃で3時間照射した。次いで、反応物を、水を添加することによってクエンチした。得られた溶液をジクロロメタン/MeOH=10:1で抽出し、水層を合わせて、真空下で濃縮した。残渣を、ジクロロメタン/メタノール(7:3)でシリカゲルカラム上に適用した。収集した画分をまとめ、真空下で濃縮した。これにより、519mg(67%)の2-(3-[4-[3-(3-アミノ-6-クロロピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル]プロポキシ)エタン-1-オールを黄色の油として得た。 A 10 mL microwave tube was charged with 2-[3-(4-[3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl)propoxy]ethan-1-ol (387 mg, 1.85 mmol, 1.00 equiv.), 4-bromo-6-chloropyridazin-3-amine (1.54 g, 7.39 mmol, 4.00 equiv.), DMSO (8 mL), and DIEA (1.53 mL, 5.00 equiv.). The final reaction mixture was irradiated with microwave radiation at 130° C. for 3 h. The reaction was then quenched by adding water. The resulting solution was extracted with dichloromethane/MeOH=10:1, and the aqueous layers were combined and concentrated under vacuum. The residue was applied onto a silica gel column with dichloromethane/methanol (7:3). The collected fractions were combined and concentrated under vacuum. This gave 519 mg (67%) of 2-(3-[4-[3-(3-amino-6-chloropyridazin-4-yl)-3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl]propoxy)ethan-1-ol as a yellow oil.
工程7

窒素不活性雰囲気下でパージし、維持した50mLの丸底フラスコに、2-(3-[4-[3-(3-アミノ-6-クロロピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル]プロポキシ)エタン-1-オール(650mg、1.55mmol、1.00当量)、[2-(メトキシメトキシ)フェニル]ボロン酸(566mg、3.11mmol、2.00当量)、ジオキサン(8mL)、水(2mL)、炭酸カリウム(644mg、4.66mmol、3.00当量)、Pd(PPh3)4(180mg、0.16mmol、0.10当量)を入れた。得られた溶液を、油浴中で、100℃で3時間撹拌した。得られた混合物を真空下で濃縮した。残渣を、メタノール/H2O(85:15)でシリカゲルカラム上に適用した。収集した画分をまとめ、真空下で濃縮した。これにより、400mg(50%)の2-[3-[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]プロポキシ]エタン-1-オールを黄色の油として得た。 A 50 mL round bottom flask purged and maintained under nitrogen inert atmosphere was charged with 2-(3-[4-[3-(3-amino-6-chloropyridazin-4-yl)-3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl]propoxy)ethan-1-ol (650 mg, 1.55 mmol, 1.00 equiv.), [2-(methoxymethoxy)phenyl]boronic acid (566 mg, 3.11 mmol, 2.00 equiv.), dioxane (8 mL), water (2 mL), potassium carbonate (644 mg, 4.66 mmol, 3.00 equiv.), and Pd(PPh3)4 (180 mg, 0.16 mmol, 0.10 equiv.). The resulting solution was stirred in an oil bath at 100 °C for 3 h. The resulting mixture was concentrated under vacuum. The residue was applied onto a silica gel column with methanol/HO (85:15). The collected fractions were combined and concentrated under vacuum. This gave 400 mg (50%) of 2-[3-[4-(3-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl]propoxy]ethan-1-ol as a yellow oil.
工程8

50mLの丸底フラスコに、2-[3-[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]プロポキシ]エタン-1-オール(400mg、0.77mmol、1.00当量)、ジクロロメタン(15mL)、TsCl(219mg、1.15mmol、1.50当量)、トリエチルアミン(155mg、1.53mmol、2.00当量)、4-ジメチルアミノピリジン(9.4mg、0.08mmol、0.10当量)を入れた。得られた溶液を、室温で一晩撹拌した。次いで、反応を、水の添加により停止させた。得られた溶液をジクロロメタンで抽出し、有機層を合わせた。得られた混合物を、飽和塩化ナトリウムで洗浄した。混合物を、無水硫酸ナトリウムで乾燥させた。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。収集した画分をまとめ、真空下で濃縮した。これにより、330mg(64%)の2-[3-[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]プロポキシ]エチル4-メチルベンゼン-1-スルホネートを黄色の固体として得た。 A 50 mL round bottom flask was charged with 2-[3-[4-(3-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl]propoxy]ethan-1-ol (400 mg, 0.77 mmol, 1.00 equiv.), dichloromethane (15 mL), TsCl (219 mg, 1.15 mmol, 1.50 equiv.), triethylamine (155 mg, 1.53 mmol, 2.00 equiv.), 4-dimethylaminopyridine (9.4 mg, 0.08 mmol, 0.10 equiv.). The resulting solution was stirred overnight at room temperature. The reaction was then quenched by the addition of water. The resulting solution was extracted with dichloromethane and the organic layers were combined. The resulting mixture was washed with saturated sodium chloride. The mixture was dried over anhydrous sodium sulfate. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (10:1). The collected fractions were combined and concentrated under vacuum. This gave 330 mg (64%) of 2-[3-[4-(3-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl]propoxy]ethyl 4-methylbenzene-1-sulfonate as a yellow solid.
工程9
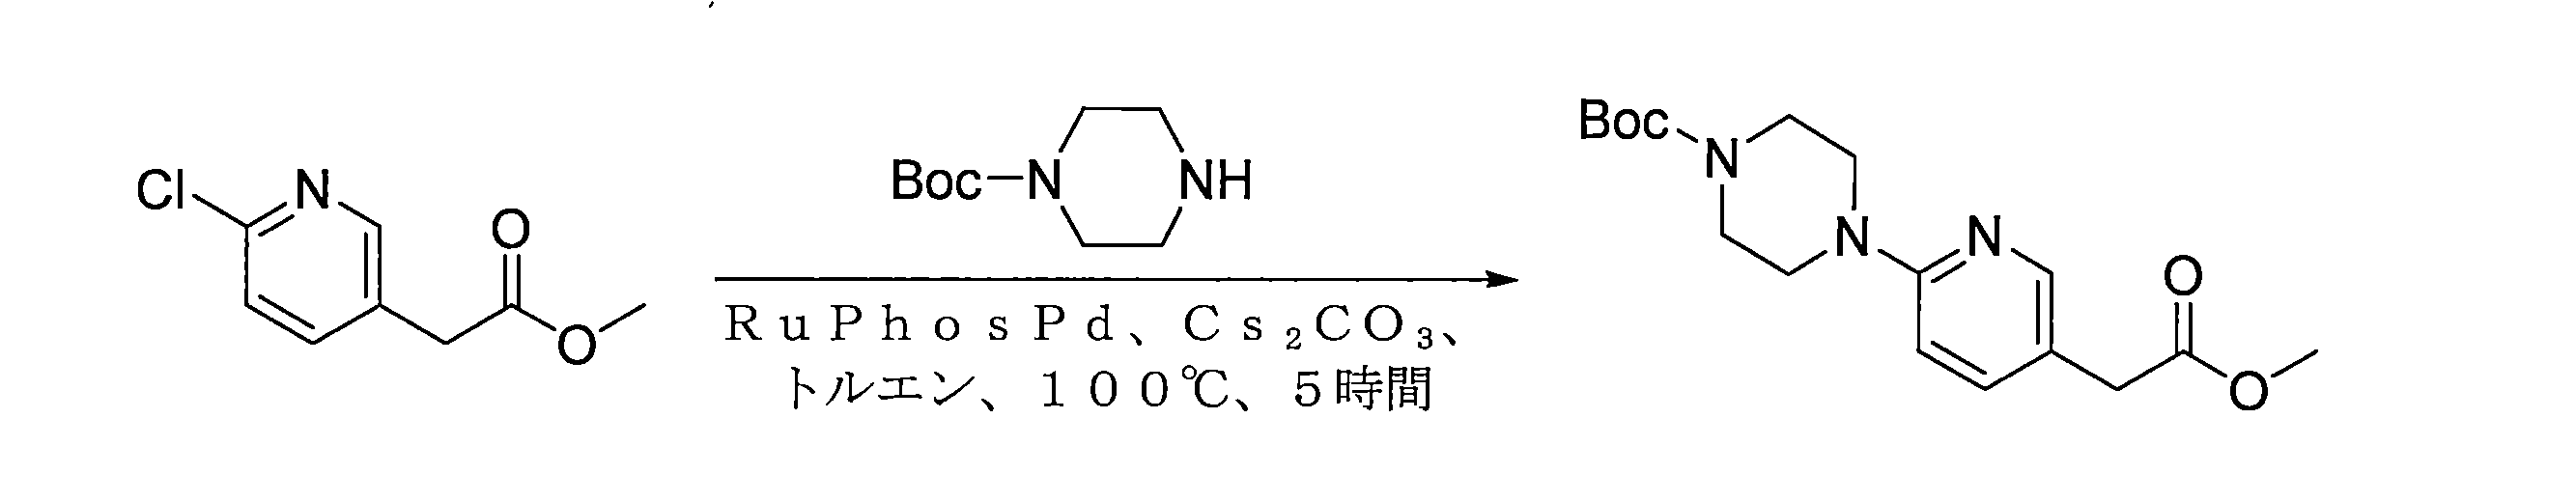
窒素不活性雰囲気下でパージし、維持した50mLの丸底フラスコに、メチル2-(6-クロロピリジン-3-イル)酢酸塩(500mg、2.69mmol、1.00当量)、tert-ブチルピペラジン-1-カルボキシレート(502mg、2.70mmol、1.00当量)、Cs2CO3(2.63g、8.07mmol、3.00当量)、トルエン(10mL)、RuPhosPd(115mg、0.05当量)を入れた。得られた溶液を一晩、油浴中、100℃で撹拌した。次いで、反応を、水の添加により停止させた。得られた溶液を酢酸エチルで抽出し、有機層を合わせた。得られた混合物を、飽和塩化ナトリウムで洗浄した。混合物を、無水硫酸ナトリウムで乾燥させた。残渣を、酢酸エチル/石油エーテル(1:1)で溶出するシリカゲルカラム上に適用した。収集した画分をまとめ、真空下で濃縮した。これにより、319mg(35%)のtert-ブチル4-[5-(2-メトキシ-2-オキソエチル)ピリジン-2-イル]ピペラジン-1-カルボキシレートを黄色の固体として得た。 A 50 mL round bottom flask purged and maintained under nitrogen inert atmosphere was charged with methyl 2-(6-chloropyridin-3-yl)acetate (500 mg, 2.69 mmol, 1.00 equiv.), tert-butyl piperazine-1-carboxylate (502 mg, 2.70 mmol, 1.00 equiv.), Cs2CO3 (2.63 g, 8.07 mmol, 3.00 equiv.), toluene (10 mL), and RuPhosPd (115 mg, 0.05 equiv.). The resulting solution was stirred overnight in an oil bath at 100°C. The reaction was then quenched by the addition of water. The resulting solution was extracted with ethyl acetate and the organic layers were combined. The resulting mixture was washed with saturated sodium chloride. The mixture was dried over anhydrous sodium sulfate. The residue was applied onto a silica gel column eluting with ethyl acetate/petroleum ether (1:1). The collected fractions were combined and concentrated under vacuum. This gave 319 mg (35%) of tert-butyl 4-[5-(2-methoxy-2-oxoethyl)pyridin-2-yl]piperazine-1-carboxylate as a yellow solid.
工程10
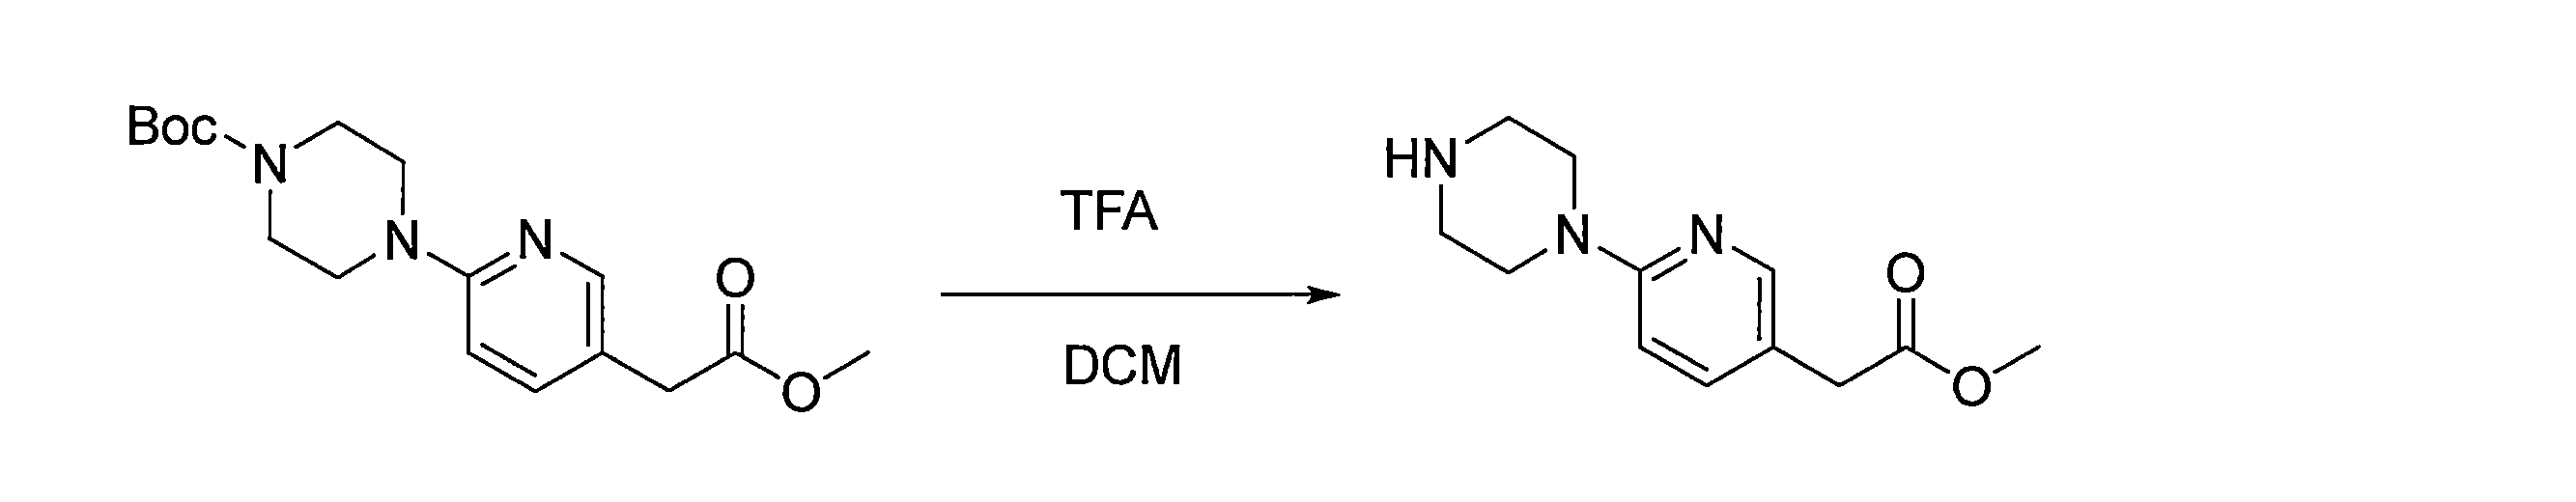
50mLの丸底フラスコに、tert-ブチル4-[5-(2-メトキシ-2-オキソエチル)ピリジン-2-イル]ピペラジン-1-カルボキシレート(319mg、0.95mmol、1.00当量)、ジクロロメタン(8mL)、トリフルオロ酢酸(2mL)を入れた。得られた溶液を室温で2時間撹拌した。得られた混合物を真空下で濃縮した。これにより、223mg(100%)のメチル2-[6-(ピペラジン-1-イル)ピリジン-3-イル]酢酸塩を黄色の油として得た。 A 50 mL round bottom flask was charged with tert-butyl 4-[5-(2-methoxy-2-oxoethyl)pyridin-2-yl]piperazine-1-carboxylate (319 mg, 0.95 mmol, 1.00 equiv.), dichloromethane (8 mL), and trifluoroacetic acid (2 mL). The resulting solution was stirred at room temperature for 2 hours. The resulting mixture was concentrated under vacuum. This afforded 223 mg (100%) of methyl 2-[6-(piperazin-1-yl)pyridin-3-yl]acetate as a yellow oil.
工程11

50mLの丸底フラスコに、メチル2-[6-(ピペラジン-1-イル)ピリジン-3-イル]酢酸塩(101mg、0.43mmol、1.00当量)、2-3-[4-(3-3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]プロポキシエチル4-メチルベンゼン-1-スルホネート(320mg、0.47mmol、1.10当量)、アセトニトリル(4mL)、炭酸カリウム(298mg、2.16mmol、5.00当量)、NaI(193mg、3.00当量)を入れた。得られた溶液を60℃で一晩撹拌した。得られた混合物を、真空下で濃縮した。残渣をジクロロメタンと水とに分割し、有機層を分離させ、飽和塩化ナトリウムで洗浄した。混合物を、無水硫酸ナトリウムで乾燥させた。残渣を、ジクロロメタン/メタノール(5:1)で溶出するシリカゲルカラム上に適用した。収集した画分をまとめ、真空下で濃縮した。これにより、94mg(30%)のメチル2-[6-[4-(2-[3-[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]プロポキシ]エチル)ピペラジン-1-イル]ピリジン-3-イル]酢酸塩を黄色の固体として得た。 A 50 mL round bottom flask was charged with methyl 2-[6-(piperazin-1-yl)pyridin-3-yl]acetate (101 mg, 0.43 mmol, 1.00 equiv.), 2-3-[4-(3-3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl]propoxyethyl 4-methylbenzene-1-sulfonate (320 mg, 0.47 mmol, 1.10 equiv.), acetonitrile (4 mL), potassium carbonate (298 mg, 2.16 mmol, 5.00 equiv.), and NaI (193 mg, 3.00 equiv.). The resulting solution was stirred at 60° C. overnight. The resulting mixture was concentrated under vacuum. The residue was partitioned between dichloromethane and water, and the organic layer was separated and washed with saturated sodium chloride. The mixture was dried over anhydrous sodium sulfate. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (5:1). The collected fractions were combined and concentrated under vacuum. This gave 94 mg (30%) of methyl 2-[6-[4-(2-[3-[4-(3-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl]propoxy]ethyl)piperazin-1-yl]pyridin-3-yl]acetate as a yellow solid.
工程12

50mLの丸底フラスコに、メチル2-[6-[4-(2-[3-[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]プロポキシ]エチル)ピペラジン-1-イル]ピリジン-3-イル]酢酸塩(94mg、0.13mmol、1.00当量)、メタノール(5mL)、水(2mL)、およびLiOH(15mg、0.64mmol、5.00当量)を入れた。得られた溶液を、室温で2日間撹拌した。得られた混合物を真空下で濃縮した。得られた溶液を10mLのH2Oで希釈した。得られた溶液をジクロロメタンで抽出し、水層を合わせて、真空下で濃縮した。残渣を、10mLのメタノール中に溶解した。固体を濾別した。得られた混合物を真空下で濃縮した。これにより、72mg(78%)の2-[6-[4-(2-[3-[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]プロポキシ]エチル)ピペラジン-1-イル]ピリジン-3-イル]酢酸を黄色の固体として得た。 A 50 mL round bottom flask was charged with methyl 2-[6-[4-(2-[3-[4-(3-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl]propoxy]ethyl)piperazin-1-yl]pyridin-3-yl]acetate (94 mg, 0.13 mmol, 1.00 equiv.), methanol (5 mL), water (2 mL), and LiOH (15 mg, 0.64 mmol, 5.00 equiv.). The resulting solution was stirred at room temperature for 2 days. The resulting mixture was concentrated under vacuum. The resulting solution was diluted with 10 mL of H2O. The resulting solution was extracted with dichloromethane and the aqueous layers were combined and concentrated under vacuum. The residue was dissolved in 10 mL of methanol. The solids were filtered off. The resulting mixture was concentrated under vacuum. This gave 72 mg (78%) of 2-[6-[4-(2-[3-[4-(3-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl]propoxy]ethyl)piperazin-1-yl]pyridin-3-yl]acetic acid as a yellow solid.
以下のスキームに従って、かつ前述の実施例について上に記載される手順を使用して、2-[6-[4-(2-[3-[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]プロポキシ]エチル)ピペラジン-1-イル]ピリジン-3-イル]酢酸を、最終化合物(2S,4R)-1-((2S)-2-(2-(6-(4-(2-(3-(4-(3-(3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル)プロポキシ)エチル)ピペラジン-1-イル)ピリジン-3-イル)アセトアミド)-3,3-ジメチルブタノイル)-4-ヒドロキシ-N-(4-(4-メチルチアゾール-5-イル)ベンジル)ピロリジン-2-カルボキサミドに変換した。
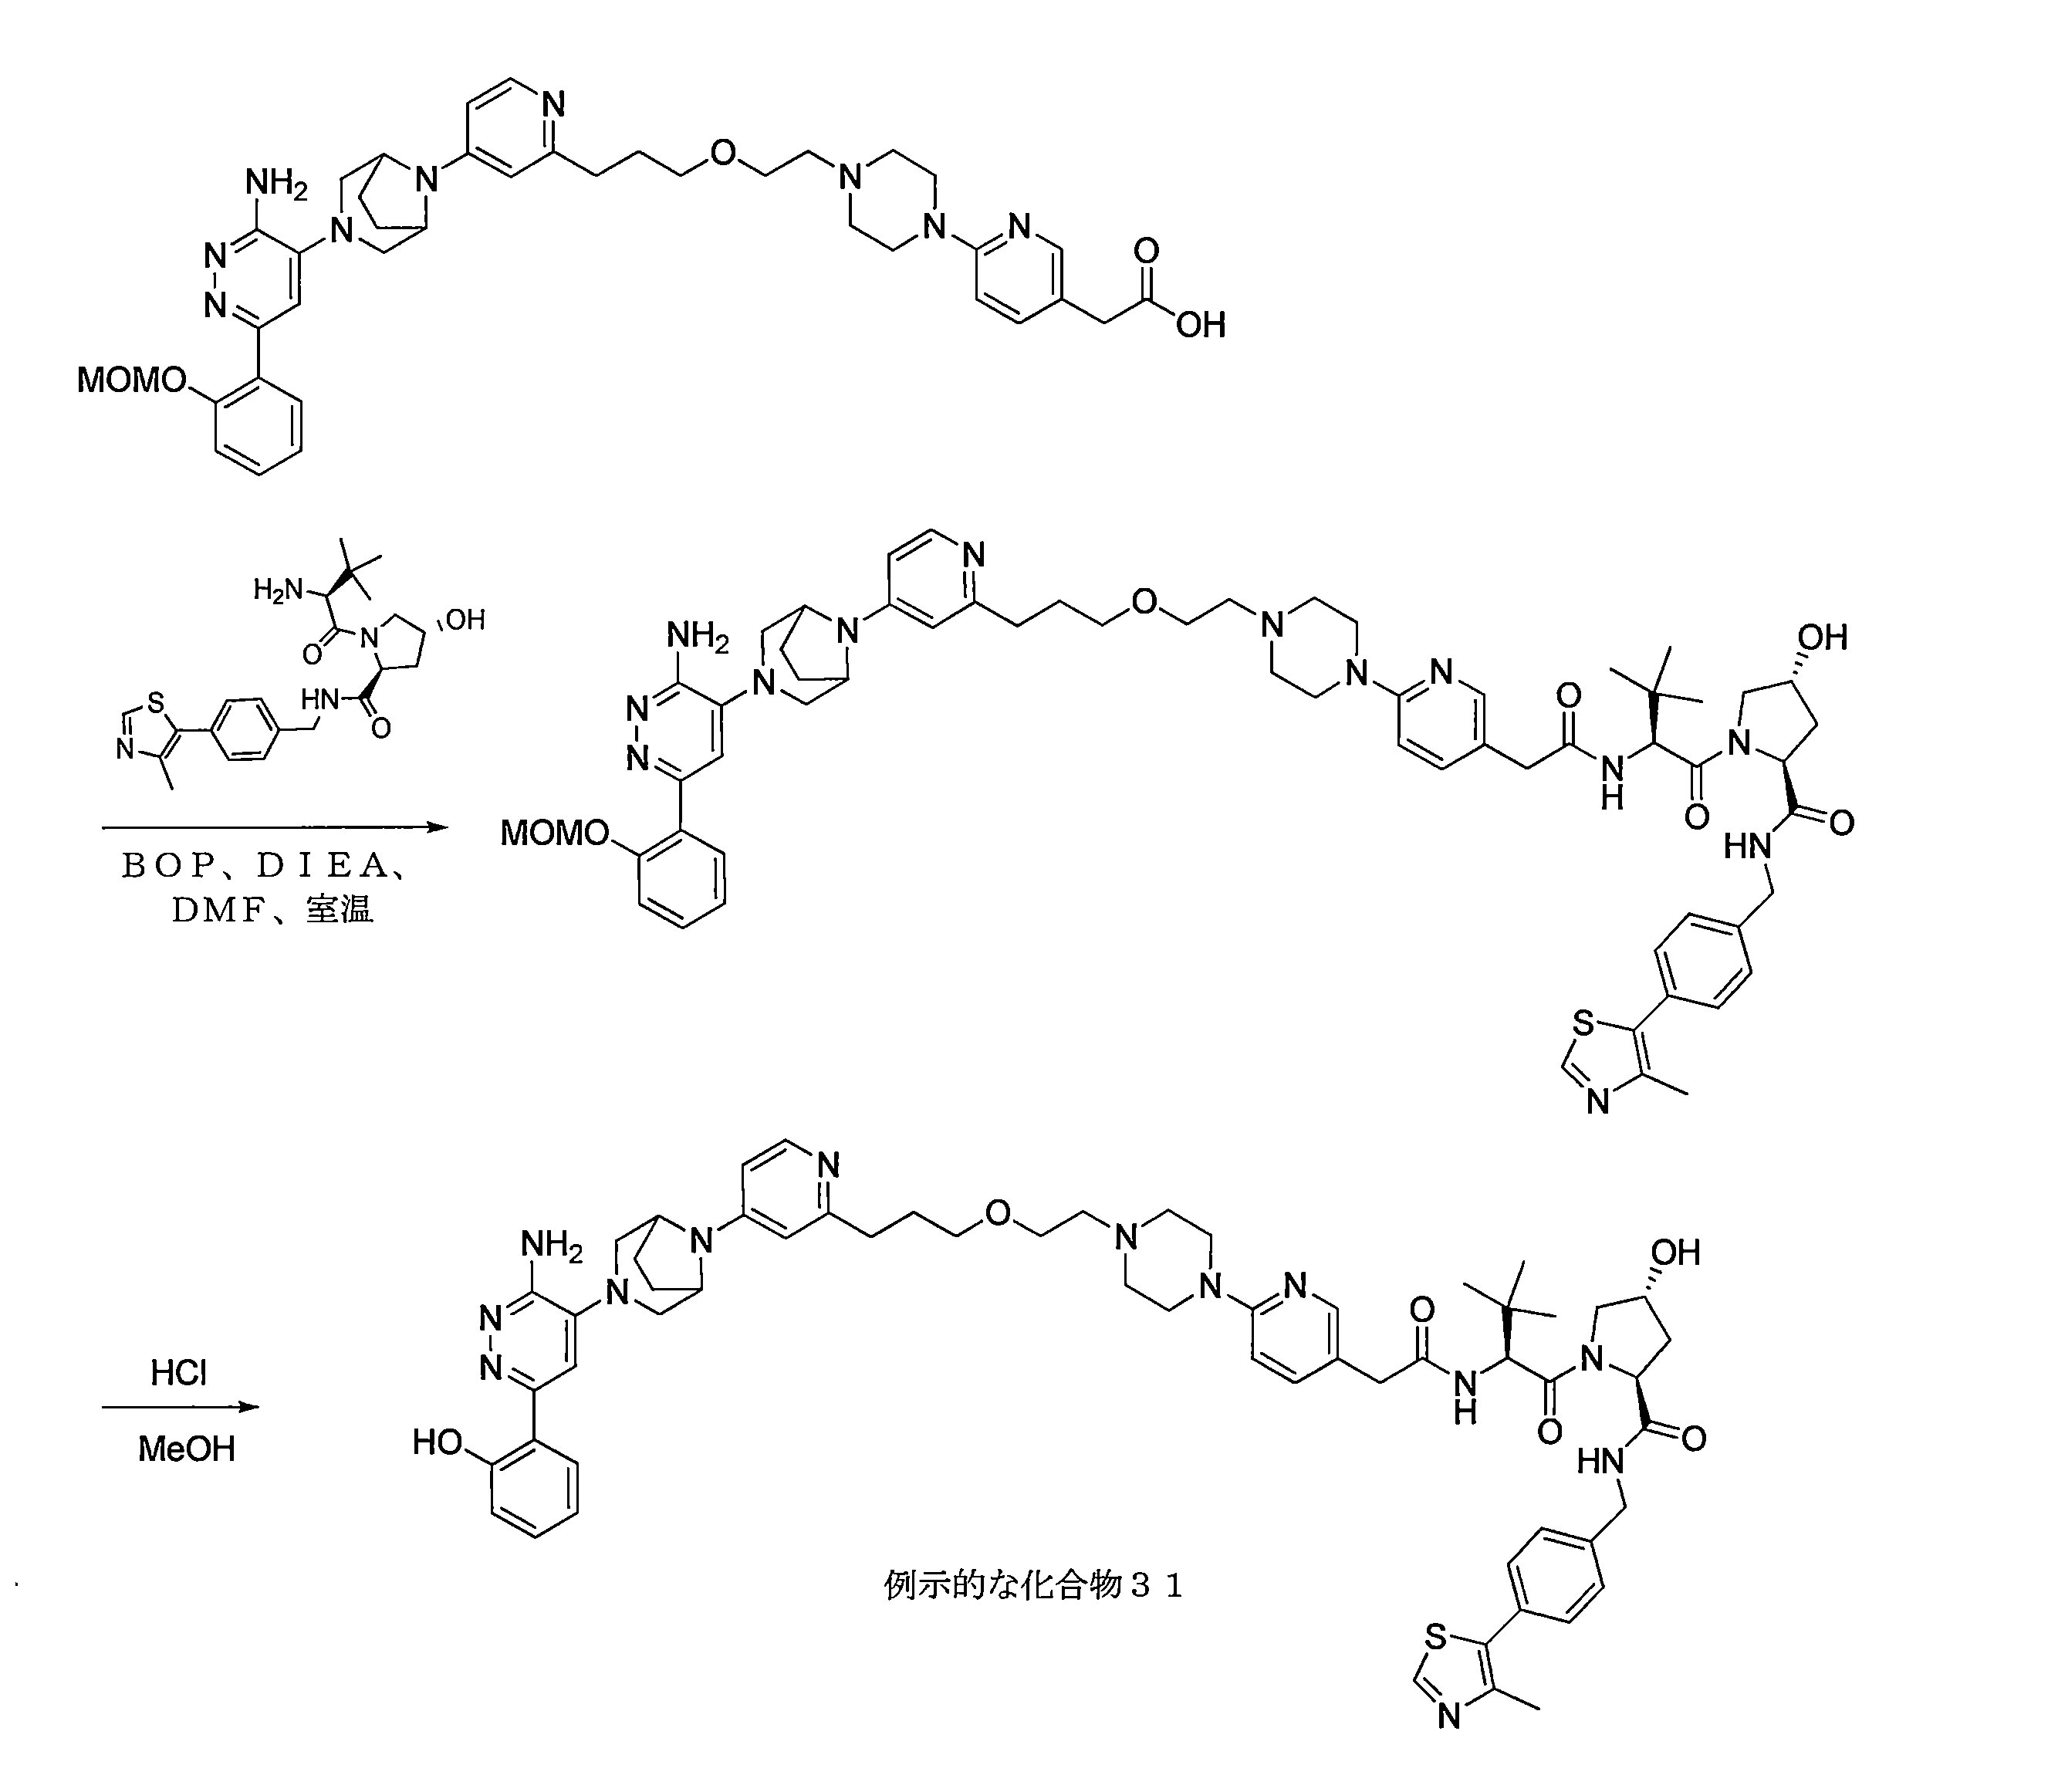
例示的な化合物50を、例示的な化合物31について記載される手順を使用して調製した。 Exemplary compound 50 was prepared using the procedure described for exemplary compound 31.
例示的な化合物32の例示的な合成
工程1

窒素不活性雰囲気下でパージし、維持した250mLの丸底フラスコに、ジクロロメタン(100mL)中の2-[2-(2-ヒドロキシエトキシ)エトキシ]エタン-1-オール(10.0g、66.59mmol、1.00当量)の溶液、(ジエチルオキソニオ)トリフルオロボレート(1.9g、13.33mmol、0.20当量)、およびエチル2-ジアゾ酢酸塩(3.8g、0.50当量)を入れた。得られた溶液を室温で3時間撹拌した。次いで、反応を、水の添加により停止させた。得られた溶液を酢酸エチルで抽出し、有機層を合わせた。得られた混合物をブラインで洗浄した。混合物を、無水硫酸ナトリウムで乾燥させた。残渣を、酢酸エチル/石油エーテル(1:1)とともにシリカゲルカラムに充填させた。収集した画分をまとめ、真空下で濃縮した。これにより、3.6g(23%)のエチル2-[2-[2-(2-ヒドロキシエトキシ)エトキシ]エトキシ]酢酸塩を黄色の油として得た。 A 250 mL round bottom flask purged and maintained under nitrogen inert atmosphere was charged with a solution of 2-[2-(2-hydroxyethoxy)ethoxy]ethan-1-ol (10.0 g, 66.59 mmol, 1.00 equiv.), (diethyloxonio)trifluoroborate (1.9 g, 13.33 mmol, 0.20 equiv.), and ethyl 2-diazoacetate (3.8 g, 0.50 equiv.) in dichloromethane (100 mL). The resulting solution was stirred at room temperature for 3 hours. The reaction was then quenched by the addition of water. The resulting solution was extracted with ethyl acetate and the organic layers were combined. The resulting mixture was washed with brine. The mixture was dried over anhydrous sodium sulfate. The residue was loaded onto a silica gel column with ethyl acetate/petroleum ether (1:1). The collected fractions were combined and concentrated under vacuum. This yielded 3.6 g (23%) of ethyl 2-[2-[2-(2-hydroxyethoxy)ethoxy]ethoxy]acetate as a yellow oil.
工程2

窒素不活性雰囲気下でパージし、維持した50mLの丸底フラスコに、ジクロロメタン(20mL)中のエチル2-[2-[2-(2-ヒドロキシエトキシ)エトキシ]エトキシ]酢酸塩(1.0g、4.23mmol、1.00当量)の溶液、4-メチルベンゼン-1-スルホニルクロリド(970mg、5.09mmol、1.20当量)、トリエチルアミン(860.0mg、8.50mmol、2.00当量)を入れた。得られた溶液を室温で5時間撹拌した。次いで、反応を、水の添加により停止させた。得られた溶液を酢酸エチルで抽出し、有機層を合わせた。得られた混合物をブラインで洗浄した。混合物を、無水硫酸ナトリウムで乾燥させた。残渣を、酢酸エチル/石油エーテル(1:1)とともにシリカゲルカラムに充填させた。収集した画分をまとめ、真空下で濃縮した。これにより、1.1g(65%)のエチル2-[2-[2-(2-[[(4-メチルベンゼン)スルホニル]オキシ]エトキシ)エトキシ]エトキシ]酢酸塩を黄色の油として得た。 A 50 mL round bottom flask purged and maintained under nitrogen inert atmosphere was charged with a solution of ethyl 2-[2-[2-(2-hydroxyethoxy)ethoxy]ethoxy]acetate (1.0 g, 4.23 mmol, 1.00 equiv.), 4-methylbenzene-1-sulfonyl chloride (970 mg, 5.09 mmol, 1.20 equiv.), triethylamine (860.0 mg, 8.50 mmol, 2.00 equiv.) in dichloromethane (20 mL). The resulting solution was stirred at room temperature for 5 hours. The reaction was then quenched by the addition of water. The resulting solution was extracted with ethyl acetate and the organic layers were combined. The resulting mixture was washed with brine. The mixture was dried over anhydrous sodium sulfate. The residue was loaded onto a silica gel column with ethyl acetate/petroleum ether (1:1). The collected fractions were combined and concentrated under vacuum. This yielded 1.1 g (65%) of ethyl 2-[2-[2-(2-[[(4-methylbenzene)sulfonyl]oxy]ethoxy)ethoxy]ethoxy]acetate as a yellow oil.
工程3
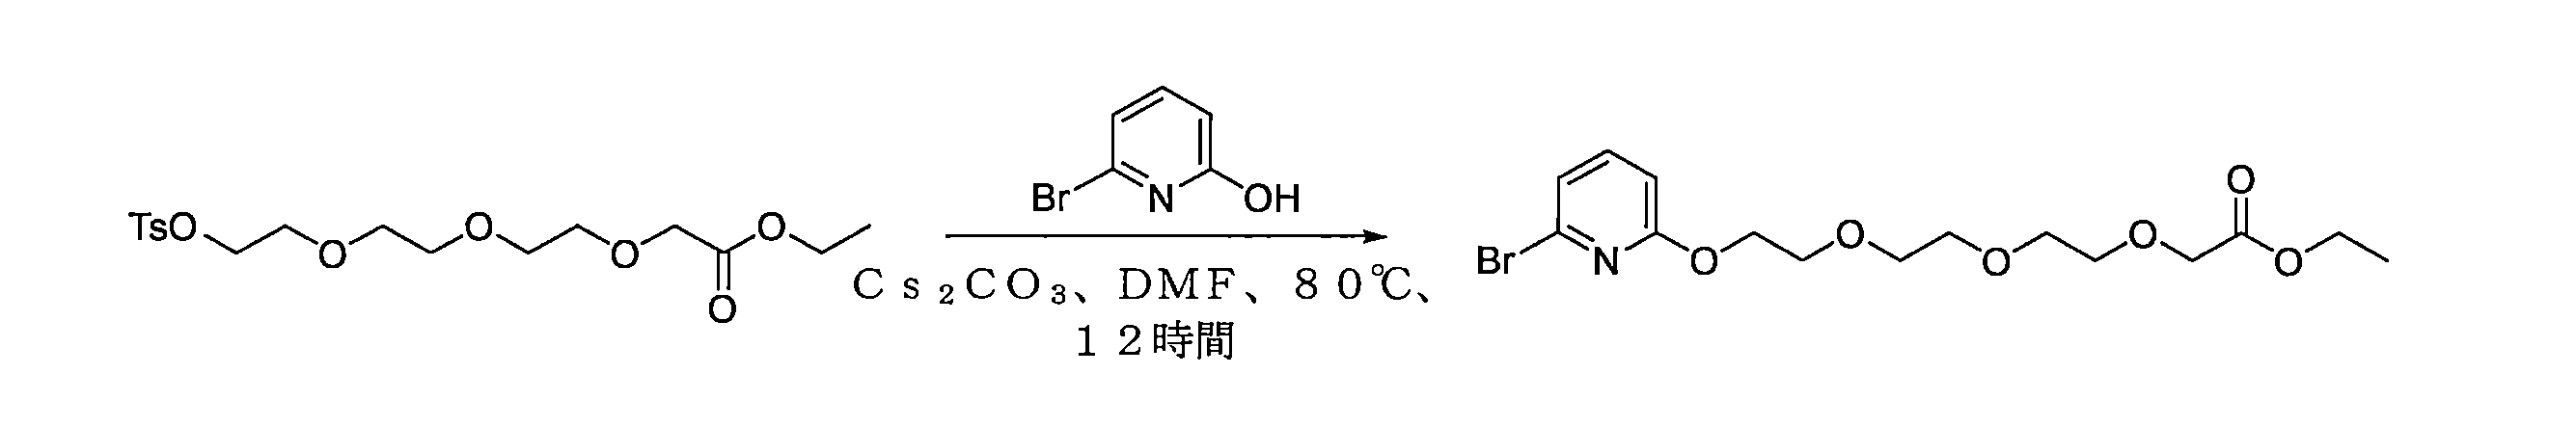
窒素不活性雰囲気下でパージし、維持した50mLの丸底フラスコに、N,N-ジメチルホルムアミド(20mL)中のエチル2-[2-[2-(2-[[(4-メチルベンゼン)スルホニル]オキシ]エトキシ)エトキシ]エトキシ]酢酸塩(1.0g、2.56mmol、1.20当量)の溶液、6-ブロモピリジン-2-オール(370mg、2.13mmol、1.00当量)、Cs2CO3(2.1g、6.45mmol、3.00当量)を入れた。得られた溶液を80℃で12時間撹拌した。次いで、反応物を、水を添加することによってクエンチした。得られた溶液を酢酸エチルで抽出し、有機層を合わせた。得られた混合物をブラインで洗浄した。混合物を、無水硫酸ナトリウムで乾燥させた。残渣を、酢酸エチル/石油エーテル(1:1)とともにシリカゲルカラムに充填させた。収集した画分をまとめ、真空下で濃縮した。これにより、800mg(96%)のエチル1-(6-ブロモピリジン-2-イル)-1,4,7,10-テトラオキサドデカン-12-エートを黄色の固体として得た。 A 50 mL round bottom flask purged and maintained under nitrogen inert atmosphere was charged with a solution of ethyl 2-[2-[2-(2-[[(4-methylbenzene)sulfonyl]oxy]ethoxy)ethoxy]ethoxy]acetate (1.0 g, 2.56 mmol, 1.20 equiv.), 6-bromopyridin-2-ol (370 mg, 2.13 mmol, 1.00 equiv.), Cs2CO3 (2.1 g, 6.45 mmol, 3.00 equiv.) in N,N-dimethylformamide (20 mL). The resulting solution was stirred at 80° C. for 12 h. The reaction was then quenched by adding water. The resulting solution was extracted with ethyl acetate and the organic layers were combined. The resulting mixture was washed with brine. The mixture was dried over anhydrous sodium sulfate. The residue was loaded onto a silica gel column with ethyl acetate/petroleum ether (1:1). The collected fractions were combined and concentrated under vacuum. This yielded 800 mg (96%) of ethyl 1-(6-bromopyridin-2-yl)-1,4,7,10-tetraoxadodecane-12-ate as a yellow solid.
以下のスキームに従って、かつ上記の他の実施例について記載されるものと類似の手順を使用して、エチル1-(6-ブロモピリジン-2-イル)-1,4,7,10-テトラオキサドデカン-12-オエートを、最終化合物(2S,4R)-1-[(2S)-2-(1-[6-[(1R,4R)-5-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-2,5-ジアザビシクロ[2.2.1ヘプタン-2-イル]ピリジン-2-イル]-1,4,7,10-テトラオキサドデカン-12-アミド)-3,3-ジメチルブタノイル]-4-ヒドロキシ-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]ピロリジン-2-カルボキサミドに変換した。

例示的な化合物33の例示的な合成
工程1

250mLの丸底フラスコに、1-ブロモ-4-エチルベンゼン(5.5g、29.8mmol、1.0当量)、CCl4(100mL)、AIBN(490mg、3.0mmol、0.1当量)、N-ブロモスクシンイミド(5.34g、30.0mmol、1.0当量)を入れた。得られた溶液を90℃で3時間撹拌した。得られた混合物を、真空下で濃縮した。これにより、5.4g(68%)の1-ブロモ-4-(1-ブロモエチル)ベンゼンを黄色の油として得た。 A 250 mL round bottom flask was charged with 1-bromo-4-ethylbenzene (5.5 g, 29.8 mmol, 1.0 equiv), CCl4 (100 mL), AIBN (490 mg, 3.0 mmol, 0.1 equiv), and N-bromosuccinimide (5.34 g, 30.0 mmol, 1.0 equiv). The resulting solution was stirred at 90°C for 3 h. The resulting mixture was concentrated under vacuum. This gave 5.4 g (68%) of 1-bromo-4-(1-bromoethyl)benzene as a yellow oil.
工程2
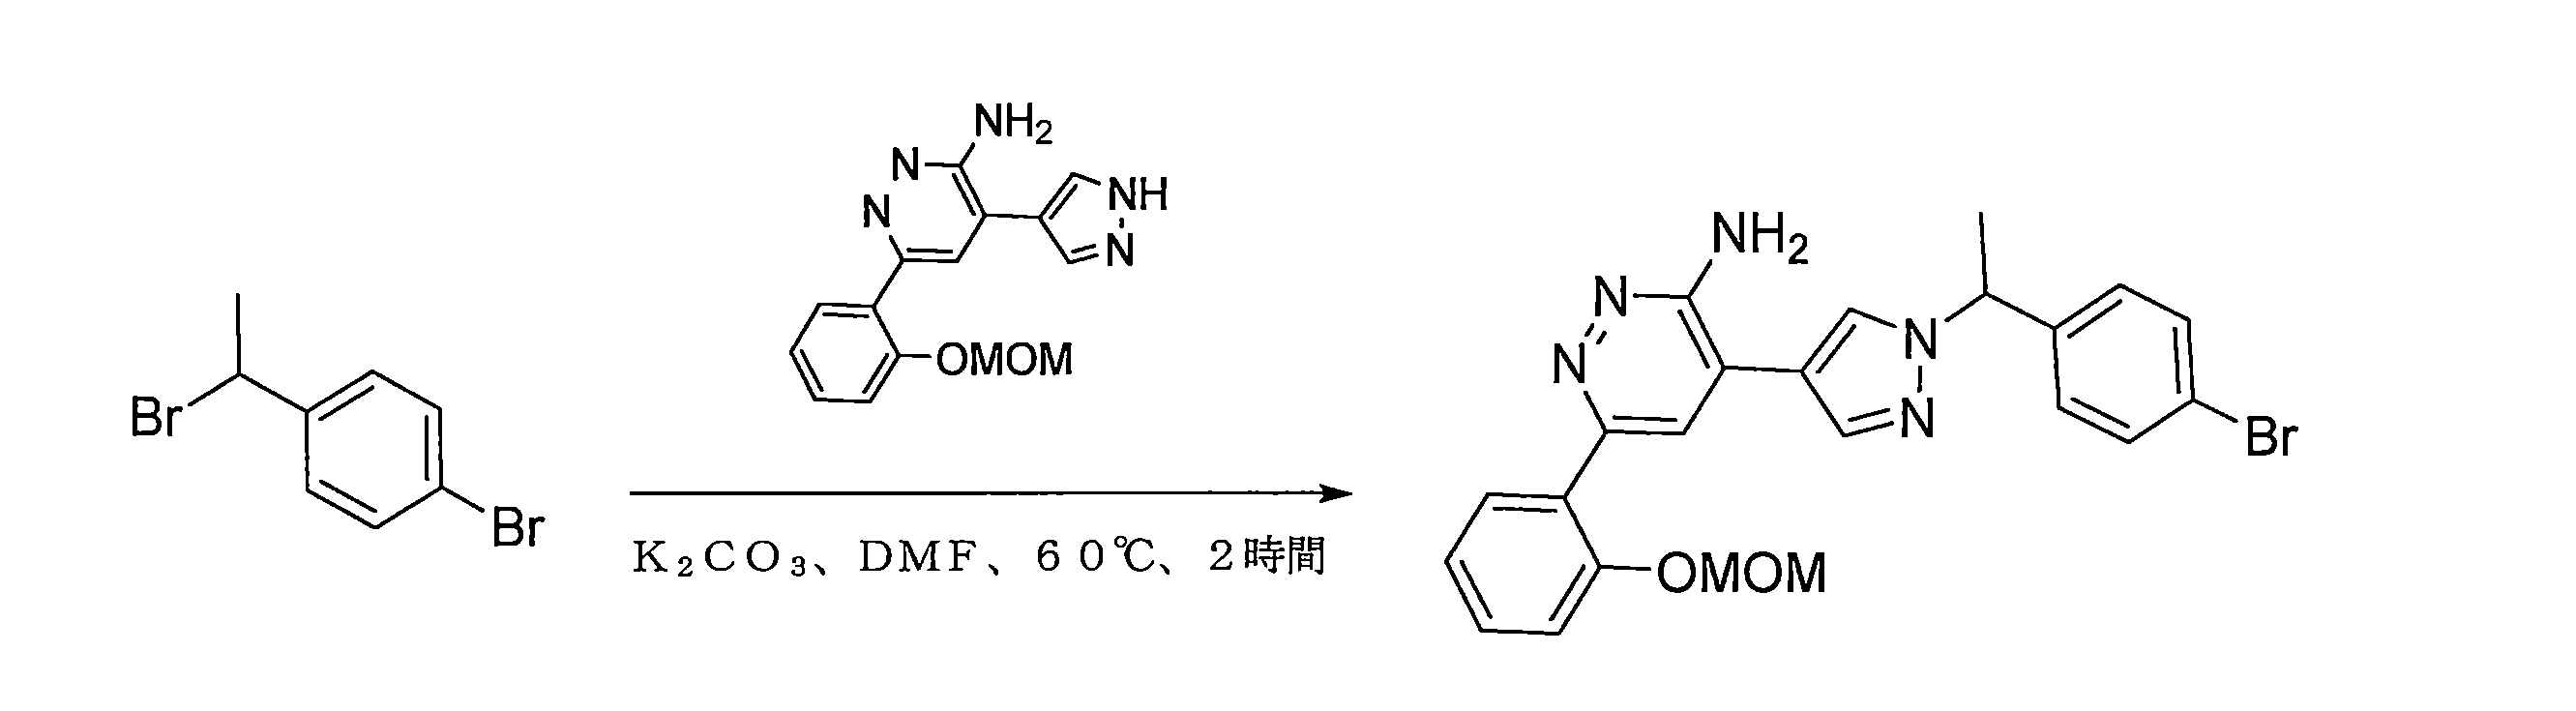
100mLの丸底フラスコに、6-[2-(メトキシメトキシ)フェニル]-4-(1H-ピラゾール-4-イル)ピリダジン-3-アミン(300mg、1.0mmol、1.0当量)、N,N-ジメチルホルムアミド(5.0mL)、1-ブロモ-4-(1-ブロモエチル)ベンゼン(400.0mg、1.5mmol、1.5当量)、炭酸カリウム(414mg、3.0mmol、3.0当量)を入れた。得られた溶液を60℃で3時間撹拌した。次いで、反応物を、10mLの水を添加することによってクエンチした。得られた溶液を酢酸エチル(20.0mL×3)で抽出し、有機層を合わせて、真空下で濃縮した。残渣を、ジクロロメタン/メタノール(1:10)で溶出するシリカゲルカラム上に適用した。これにより、300mg(62%)の4-[1-[1-(4-ブロモフェニル)エチル]-1H-ピラゾール-4-イル]-6-[2-(メトキシメトキシ)フェニル]ピリダジン-3-アミンを茶色の固体として得た。 A 100 mL round bottom flask was charged with 6-[2-(methoxymethoxy)phenyl]-4-(1H-pyrazol-4-yl)pyridazin-3-amine (300 mg, 1.0 mmol, 1.0 equiv.), N,N-dimethylformamide (5.0 mL), 1-bromo-4-(1-bromoethyl)benzene (400.0 mg, 1.5 mmol, 1.5 equiv.), and potassium carbonate (414 mg, 3.0 mmol, 3.0 equiv.). The resulting solution was stirred at 60° C. for 3 h. The reaction was then quenched by adding 10 mL of water. The resulting solution was extracted with ethyl acetate (20.0 mL×3) and the organic layers were combined and concentrated under vacuum. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (1:10). This gave 300 mg (62%) of 4-[1-[1-(4-bromophenyl)ethyl]-1H-pyrazol-4-yl]-6-[2-(methoxymethoxy)phenyl]pyridazin-3-amine as a brown solid.
工程3
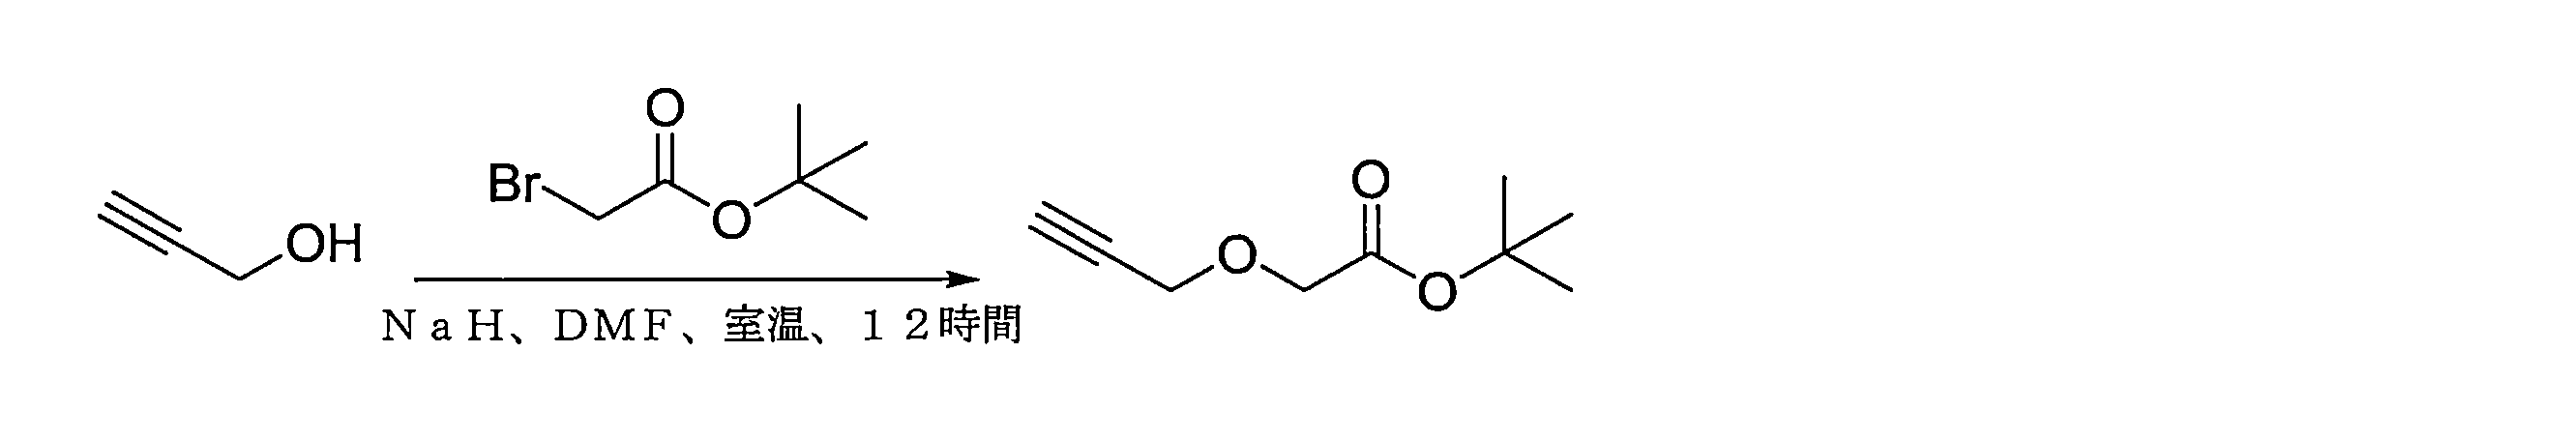
250mLの丸底フラスコに、プロパ-2-イン-1-オール(10g、178.4mmol、1.0当量)、テトラヒドロフラン(100.0mL)、水素化ナトリウム(6.4g、266.7mmol、0.9当量)、およびtert-ブチル2-ブロモ酢酸塩(28g、143.6mmol、0.8当量)を入れた。得られた溶液を室温で3時間撹拌した。次いで、反応物を、20mLの塩化アンモニウム水溶液を添加することによってクエンチした。得られた混合物を真空下で濃縮した。得られた溶液を酢酸エチル(30mL×3)で抽出し、有機層を合わせて、真空下で濃縮した。これにより、24.0g(90%)のtert-ブチル2-(プロパ-2-イン-1-イルオキシ)酢酸塩を黄色の固体として得た。 A 250 mL round bottom flask was charged with prop-2-yn-1-ol (10 g, 178.4 mmol, 1.0 equiv), tetrahydrofuran (100.0 mL), sodium hydride (6.4 g, 266.7 mmol, 0.9 equiv), and tert-butyl 2-bromoacetate (28 g, 143.6 mmol, 0.8 equiv). The resulting solution was stirred at room temperature for 3 h. The reaction was then quenched by adding 20 mL of aqueous ammonium chloride solution. The resulting mixture was concentrated under vacuum. The resulting solution was extracted with ethyl acetate (30 mL x 3), and the organic layers were combined and concentrated under vacuum. This afforded 24.0 g (90%) of tert-butyl 2-(prop-2-yn-1-yloxy)acetate as a yellow solid.
工程4

250mLの丸底フラスコに、tert-ブチル2-(プロパ-2-イン-1-イルオキシ)酢酸塩(6.6g、38.8mmol、1.0当量)、トリエチルアミン(400.0mg、3.9mmol、0.1当量)、ピナコールボラン(20.0mL)、ZrCp2HCl(1g、0.1当量)を入れた。得られた溶液を60℃で12分間撹拌した。次いで、反応物を、10mLの氷/水を添加することによってクエンチした。得られた溶液を酢酸エチル(30mL×3)で抽出し、有機層を合わせた。残渣を、酢酸エチル/石油エーテル(1:5)で溶出するシリカゲルカラム上に適用した。これにより、5.0g(43%)のtert-ブチル2-[[(2E)-3-(テトラメチル-1,3,2-ジオキサボロラン-2-イル)プロパ-2-エン-1-イル]オキシ]酢酸塩を黄色の油として得た。 A 250 mL round bottom flask was charged with tert-butyl 2-(prop-2-yn-1-yloxy)acetate (6.6 g, 38.8 mmol, 1.0 equiv), triethylamine (400.0 mg, 3.9 mmol, 0.1 equiv), pinacolborane (20.0 mL), and ZrCp2HCl (1 g, 0.1 equiv). The resulting solution was stirred at 60°C for 12 minutes. The reaction was then quenched by adding 10 mL of ice/water. The resulting solution was extracted with ethyl acetate (30 mL x 3) and the organic layers were combined. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (1:5). This gave 5.0 g (43%) of tert-butyl 2-[[(2E)-3-(tetramethyl-1,3,2-dioxaborolan-2-yl)prop-2-en-1-yl]oxy]acetate as a yellow oil.
工程5
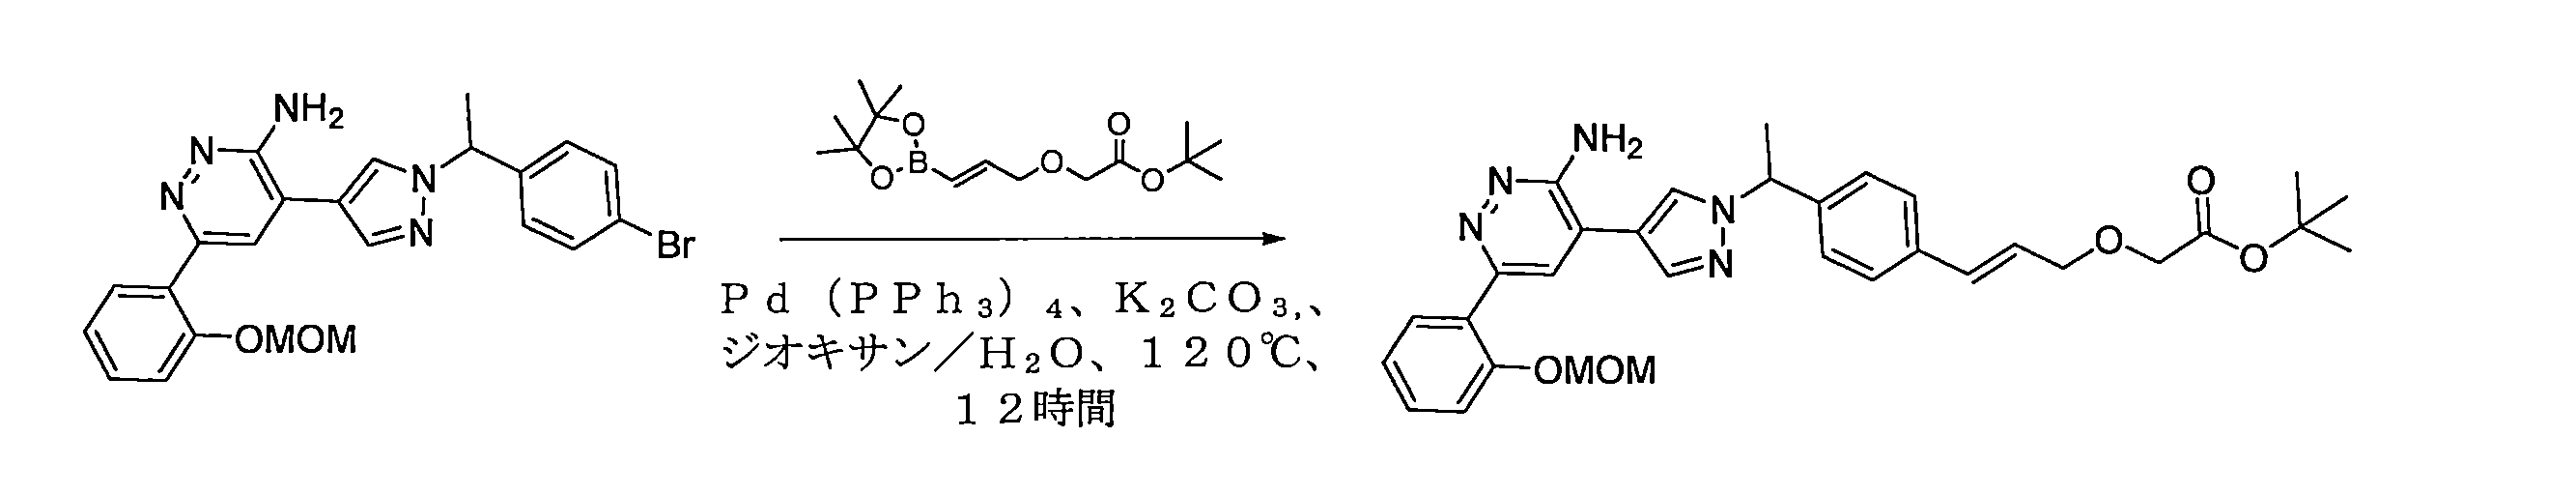
10mLの密封チューブに、4-[1-[1-(4-ブロモフェニル)エチル]-1H-ピラゾール-4-イル]-6-[2-(メトキシメトキシ)フェニル]ピリダジン-3-アミン(300mg、0.6mmol、1.0当量)、tert-ブチル2-[[(2E)-3-(テトラメチル-1,3,2-ジオキサボロラン-2-イル)プロパ-2-エン-1-イル]オキシ]酢酸塩(279mg、0.9mmol、1.5当量)、Pd(PPh3)4(72mg、0.06mmol、0.1当量)、炭酸カリウム(259mg、1.9mmol、3.0当量)、ジオキサン(4.0mL)、およびH2O(1.0mL)を入れた。得られた溶液を90℃で5時間撹拌した。得られた混合物を、真空下で濃縮した。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。これにより、200mg(56%)のtert-ブチル2-[[(2E)-3-[4-[1-(4-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-1H-ピラゾール-1-イル)エチル]フェニル]プロパ-2-エン-1-イル]オキシ]酢酸塩を黄色の固体として得た。 A 10 mL sealed tube was charged with 4-[1-[1-(4-bromophenyl)ethyl]-1H-pyrazol-4-yl]-6-[2-(methoxymethoxy)phenyl]pyridazin-3-amine (300 mg, 0.6 mmol, 1.0 equiv.), tert-butyl 2-[[(2E)-3-(tetramethyl-1,3,2-dioxaborolan-2-yl)prop-2-en-1-yl]oxy]acetate (279 mg, 0.9 mmol, 1.5 equiv.), Pd(PPh3)4 (72 mg, 0.06 mmol, 0.1 equiv.), potassium carbonate (259 mg, 1.9 mmol, 3.0 equiv.), dioxane (4.0 mL), and H2O (1.0 mL). The resulting solution was stirred at 90 °C for 5 h. The resulting mixture was concentrated under vacuum. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (10:1). This gave 200 mg (56%) of tert-butyl 2-[[(2E)-3-[4-[1-(4-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-1H-pyrazol-1-yl)ethyl]phenyl]prop-2-en-1-yl]oxy]acetate as a yellow solid.
工程6

100mLの丸底フラスコに、tert-ブチル2-[[(2E)-3-[4-[1-(4-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-1H-ピラゾール-1-イル)エチル]フェニル]プロパ-2-エン-1-イル]オキシ]酢酸塩(93mg、0.2mmol、1.0当量)、メタノール(5.0mL)、炭素上のパラジウム(100mg)を入れた。得られた溶液を、水素雰囲気下で、室温で1時間撹拌した。固形分を濾過した。得られた混合物を真空下で濃縮した。これにより、56mg(60%)のtert-ブチル2-(3-[4-[1-(4-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-1H-ピラゾール-1-イル)エチル]フェニル]プロポキシ)酢酸塩を黄色の固体として得た。 A 100 mL round bottom flask was charged with tert-butyl 2-[[(2E)-3-[4-[1-(4-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-1H-pyrazol-1-yl)ethyl]phenyl]prop-2-en-1-yl]oxy]acetate (93 mg, 0.2 mmol, 1.0 equiv.), methanol (5.0 mL), and palladium on carbon (100 mg). The resulting solution was stirred under hydrogen atmosphere at room temperature for 1 hour. The solids were filtered off. The resulting mixture was concentrated under vacuum. This afforded 56 mg (60%) of tert-butyl 2-(3-[4-[1-(4-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-1H-pyrazol-1-yl)ethyl]phenyl]propoxy)acetate as a yellow solid.
工程7

100mLの丸底フラスコに、tert-ブチル2-(3-[4-[1-(4-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-1H-ピラゾール-1-イル)エチル]フェニル]プロポキシ)酢酸塩(56mg、0.1mmol、1.0当量)、ジクロロメタン(10.0mg)、トリフルオロ酢酸(5mL)を入れた。得られた溶液を室温で2時間撹拌した。得られた混合物を真空下で濃縮した。これにより、45mg(90%)の2-(3-[4-[1-(4-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-1H-ピラゾール-1-イル)エチル]フェニル]プロポキシ)酢酸を茶色の固体として得た。 A 100 mL round bottom flask was charged with tert-butyl 2-(3-[4-[1-(4-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-1H-pyrazol-1-yl)ethyl]phenyl]propoxy)acetate (56 mg, 0.1 mmol, 1.0 equiv.), dichloromethane (10.0 mg), and trifluoroacetic acid (5 mL). The resulting solution was stirred at room temperature for 2 hours. The resulting mixture was concentrated under vacuum. This afforded 45 mg (90%) of 2-(3-[4-[1-(4-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-1H-pyrazol-1-yl)ethyl]phenyl]propoxy)acetic acid as a brown solid.
工程8

25mLの丸底フラスコに、2-[3-[4-(1-[4-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-1H-ピラゾール-1-イル]エチル)フェニル]プロポキシ]酢酸(26mg、0.06mmol、1.0当量)、N,N-ジメチルホルムアミド(5mL)、(2S,4R)-1-[(2S)-2-アミノ-3,3-ジメチルブタノイル]-4-ヒドロキシ-N-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチルピロリジン-2-カルボキサミド(30mg、0.07mmol、1.2当量)、DIEA(21mg、0.2mmol、3.0当量)、T3P(53mg、1.2当量)を入れた。得られた溶液を室温で1時間撹拌した。次いで、反応物を、0.5mLの水を添加することによってクエンチした。固体を濾別した。粗生成物(5mL)を、以下の条件を用いて、分取HPLCによって精製した。カラム、XBridge Prep C18 OBDカラム、150mm 5um;移動相、水(10MMOL/Lの重炭酸アミン)およびACN(45.0%ACN、7分間で最大52.0%);検出器、UV254/220nm。これにより、6mg(12%)の(2S,4R)-1-[(2S)-2-(2-[3-[4-(1-[4-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-1H-ピラゾール-1-イル]エチル)フェニル]プロポキシ]アセトアミド)-3,3-ジメチルブタノイル]-4-ヒドロキシ-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]ピロリジン-2-カルボキサミドを灰色の固体として得た。 A 25 mL round bottom flask was charged with 2-[3-[4-(1-[4-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-1H-pyrazol-1-yl]ethyl)phenyl]propoxy]acetic acid (26 mg, 0.06 mmol, 1.0 equiv.), N,N-dimethylformamide (5 mL), (2S,4R)-1-[(2S)-2-amino-3,3-dimethylbutanoyl]-4-hydroxy-N-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methylpyrrolidine-2-carboxamide (30 mg, 0.07 mmol, 1.2 equiv.), DIEA (21 mg, 0.2 mmol, 3.0 equiv.), and T3P (53 mg, 1.2 equiv.). The resulting solution was stirred at room temperature for 1 h. The reaction was then quenched by adding 0.5 mL of water. The solid was filtered off. The crude product (5 mL) was purified by preparative HPLC using the following conditions: column, XBridge Prep C18 OBD column, 150 mm 5 um; mobile phase, water (10 MMOL/L amine bicarbonate) and ACN (45.0% ACN, max 52.0% in 7 min); detector, UV 254/220 nm. This gave 6 mg (12%) of (2S,4R)-1-[(2S)-2-(2-[3-[4-(1-[4-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-1H-pyrazol-1-yl]ethyl)phenyl]propoxy]acetamido)-3,3-dimethylbutanoyl]-4-hydroxy-N-[[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]pyrrolidine-2-carboxamide as a grey solid.
1H NMR(300MHz、メタノール-d4)δ8.79(s、1H)、8.35~8.23(m、1H)、8.09~7.97(m、2H)、7.80(d、J=7.7Hz、1H)、7.43~7.37(m、2H)、7.31(d、J=8.1Hz、2H)、7.24(d、J=8.1Hz、1H)、7.20(s、4H)、6.89(t、J=7.6Hz、2H)、5.64~5.52(m、1H)、4.66(s、1H)、4.60~4.44(m、3H)、4.27(d、J=15.5Hz、1H)、3.92(d、J=4.6Hz、2H)、3.88~3.74(m、2H)、3.52(d、J=6.2Hz、2H)、2.71(t、J=7.6Hz、2H)、2.37(s、3H)、2.18(d、J=7.7Hz、1H)、2.07(dd、J=9.4、4.2Hz、1H)、1.89(dd、J=7.3、1.2Hz、6H)、1.26(s、1H)、1.10~0.95(d、J=1.6Hz、9H)、0.95~0.90(m、1H)。 1H NMR (300MHz, methanol-d4) δ 8.79 (s, 1H), 8.35-8.23 (m, 1H), 8.09-7.97 (m, 2H), 7.80 (d, J = 7.7 Hz, 1H), 7.43-7.37 (m, 2H), 7.31 (d, J = 8.1 Hz, 2H), 7.24 (d, J = 8.1 Hz, 1H ), 7.20 (s, 4H), 6.89 (t, J = 7.6 Hz, 2H), 5.64-5.52 (m, 1H), 4.66 (s, 1H), 4. 60-4.44 (m, 3H), 4.27 (d , J = 15.5 Hz, 1H), 3.92 (d, J = 4.6 Hz, 2H), 3.88-3.74 (m, 2H), 3.52 (d, J = 6.2 Hz, 2H), 2.71 (t, J = 7.6 Hz, 2H), 2.37 (s, 3H), 2.18 (d, J = 7.7 Hz, 1H), 2.07 (dd, J = 9.4, 4.2 Hz, 1H), 1.89 (dd, J = 7.3, 1.2 Hz, 6H), 1.26 (s, 1H), 1.10-0.95 (d, J = 1.6Hz, 9H), 0.95-0.90 (m, 1H).
例示的な化合物34の例示的な合成
例示的な化合物34を、上記の他の実施例について記載される手順、ならびに当業者に既知であり、かつ理解される手順を使用して、以下のスキームに従って調製した。
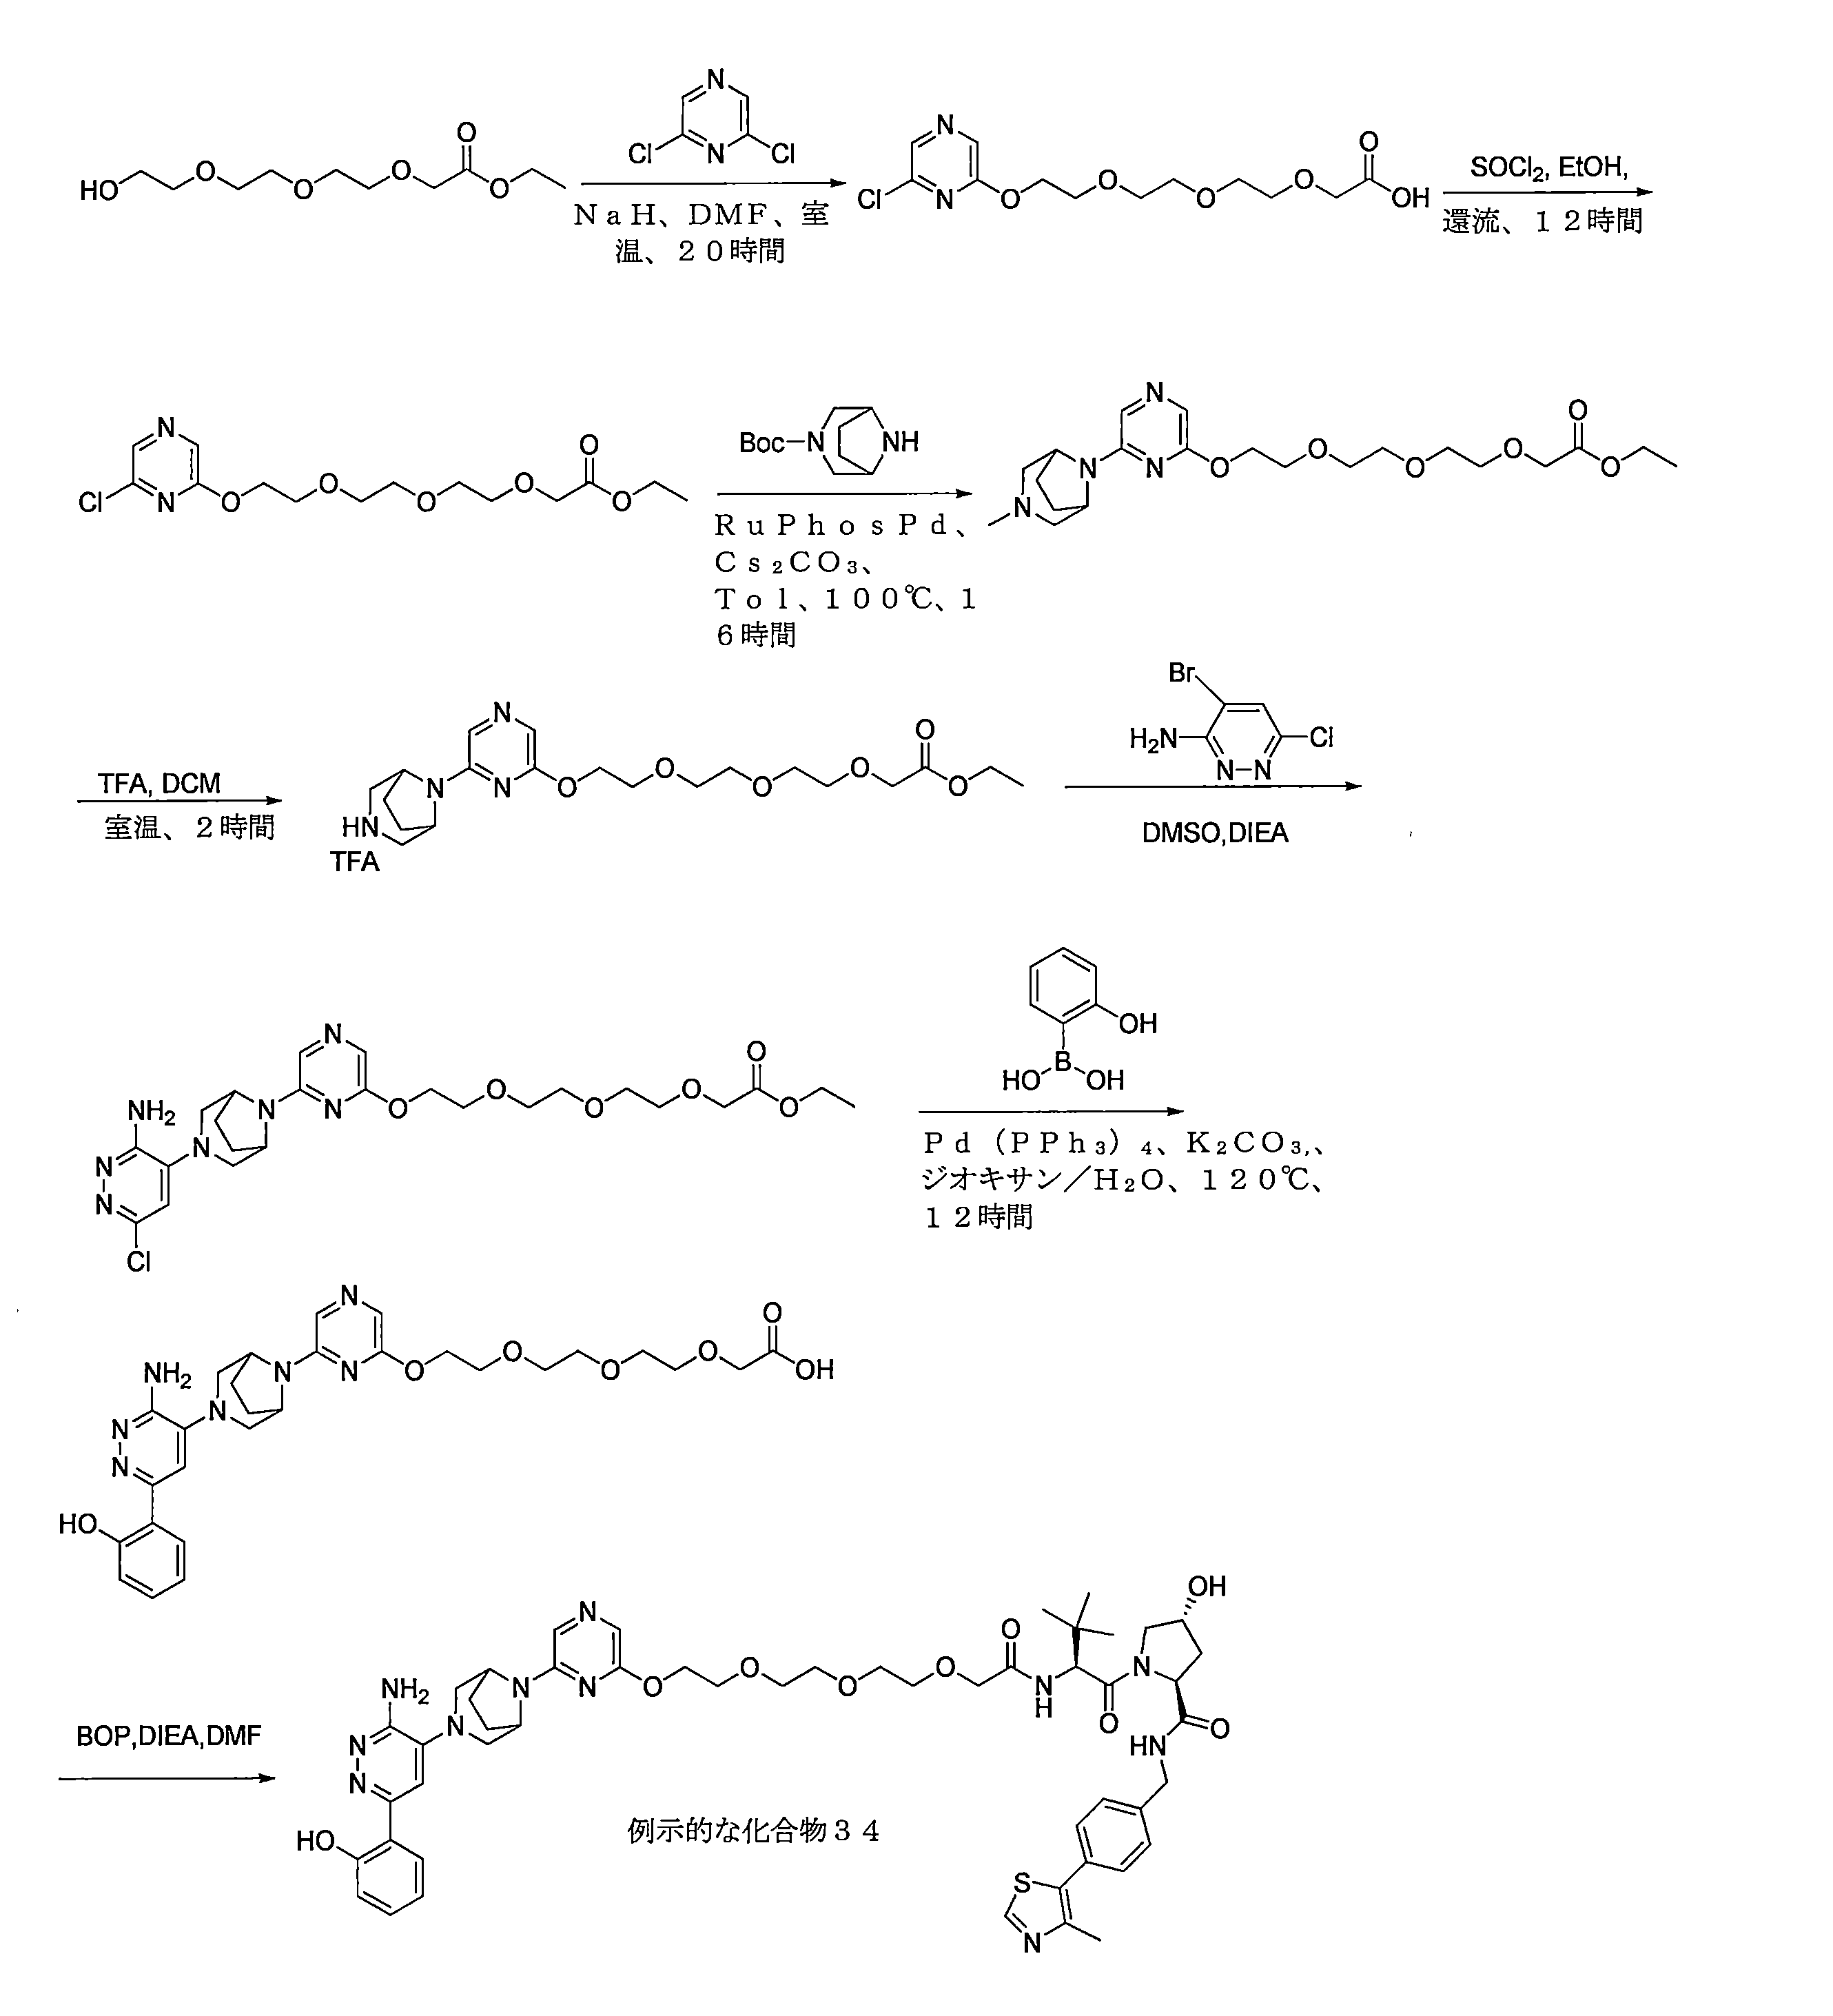
例示的な化合物36および例示的な化合物37の例示的な合成
例示的な化合物36および37を、上記の他の実施例について記載される手順、ならびに当業者に既知であり、かつ理解される手順を使用して、以下のスキームに従って調製した。
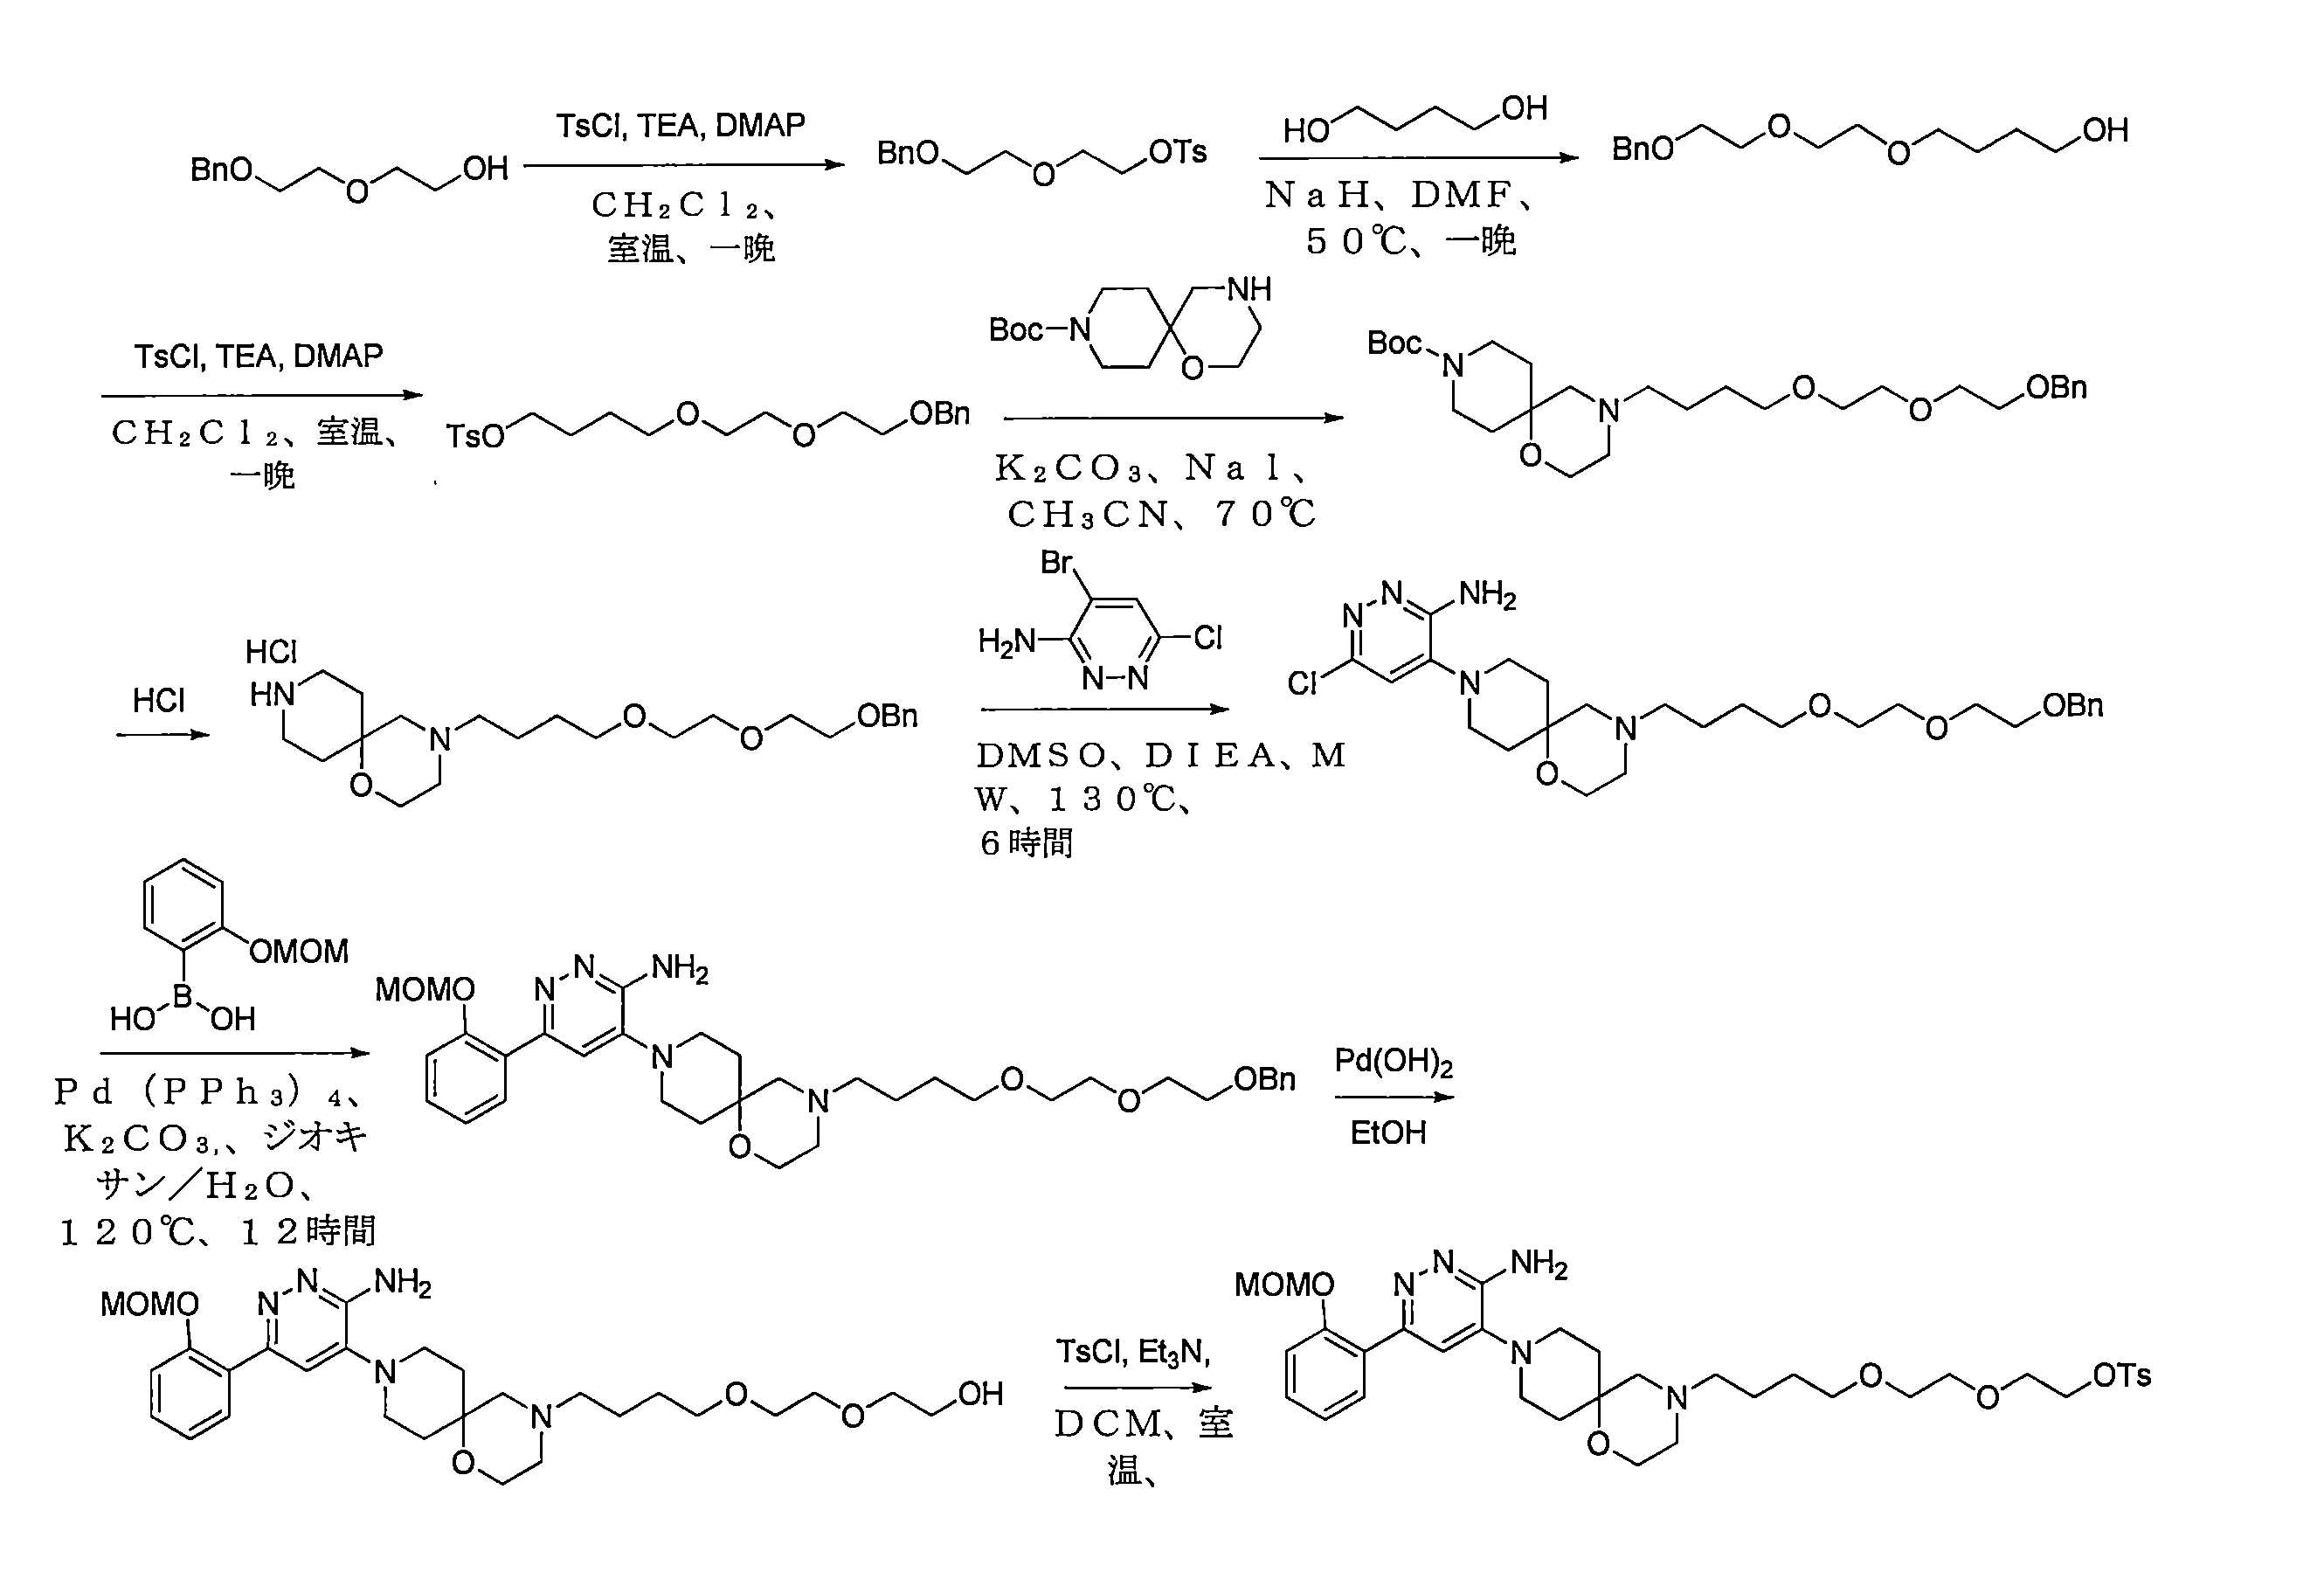

例示的な化合物38および例示的な化合物39の例示的な合成
工程1

500mLの丸底フラスコに、2-(ピペラジン-1-イル)エタン-1-オール(26.0g、199.7mmol、1.0当量)、ジクロロメタン(200.0mL)、トリエチルアミン(40.4g、399.3mmol、2.0当量)、ベンジルカルボノクロリデート(40.8g、239.2mmol、1.2当量)を入れた。得られた溶液を0℃で2時間撹拌した。次いで、反応物を、10mLの水を添加することによってクエンチした。得られた溶液をジクロロメタン(100mL×3)で抽出し、有機層を合わせて、真空下で濃縮した。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。これにより、22g(42%)のベンジル4-(2-ヒドロキシエチル)ピペラジン-1-カルボキシレートを黄色の油として得た。 A 500 mL round bottom flask was charged with 2-(piperazin-1-yl)ethan-1-ol (26.0 g, 199.7 mmol, 1.0 equiv), dichloromethane (200.0 mL), triethylamine (40.4 g, 399.3 mmol, 2.0 equiv), and benzyl carbonochloridate (40.8 g, 239.2 mmol, 1.2 equiv). The resulting solution was stirred at 0°C for 2 h. The reaction was then quenched by adding 10 mL of water. The resulting solution was extracted with dichloromethane (100 mL x 3) and the organic layers were combined and concentrated under vacuum. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (10:1). This gave 22 g (42%) of benzyl 4-(2-hydroxyethyl)piperazine-1-carboxylate as a yellow oil.
工程2
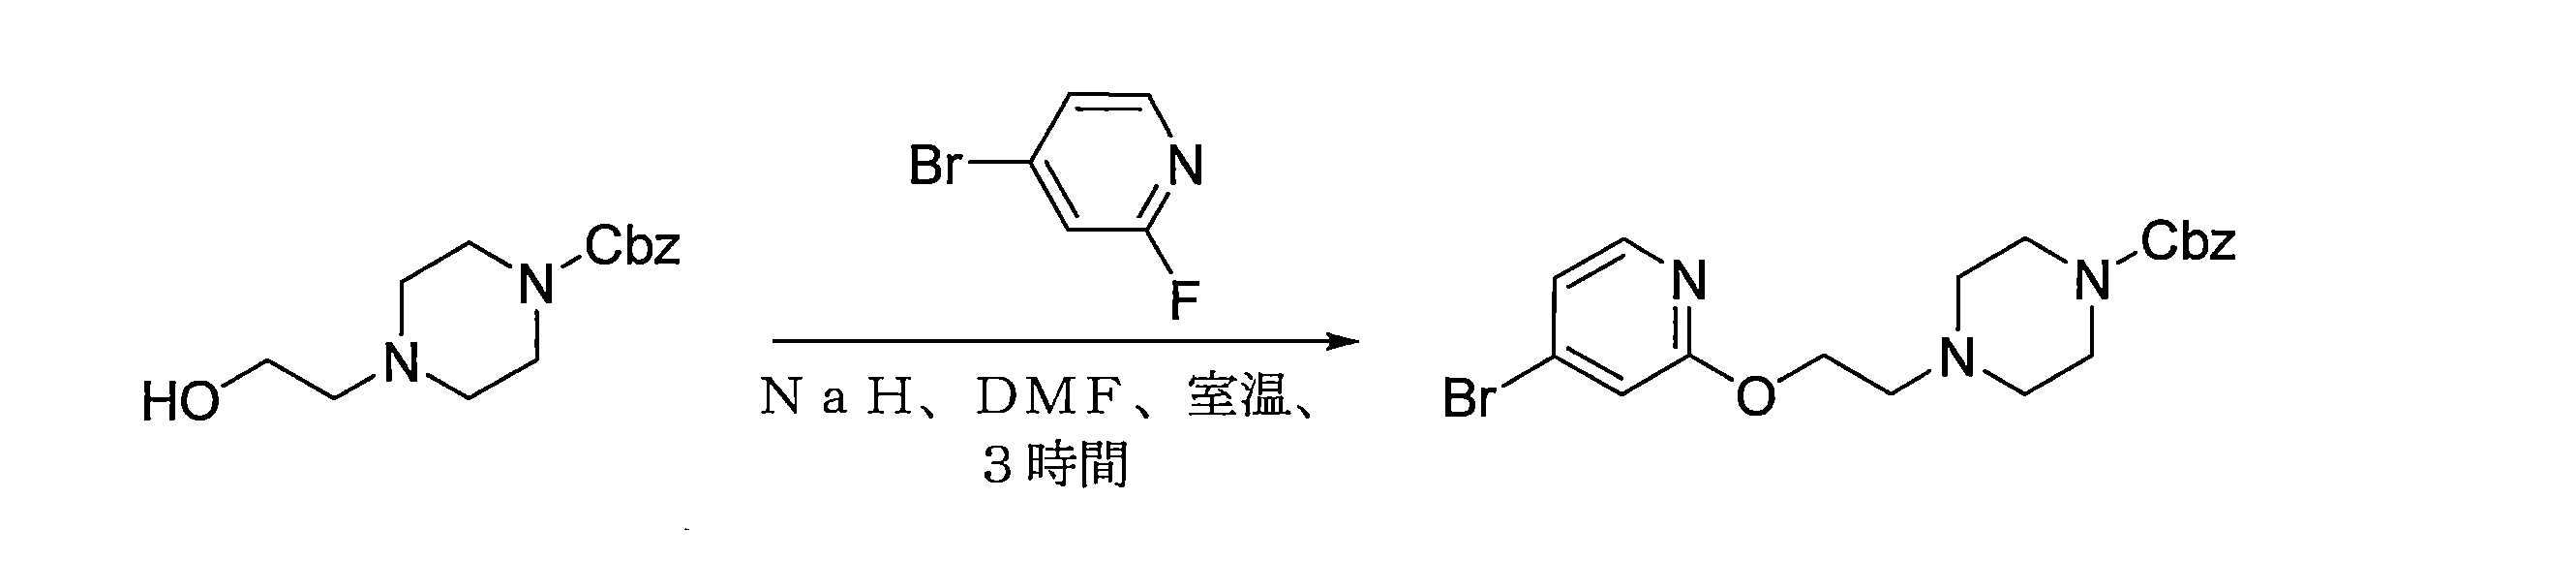
100mLの丸底フラスコに、ベンジル4-(2-ヒドロキシエチル)ピペラジン-1-カルボキシレート(5.3g、20.1mmol、1.0当量)およびN,N-ジメチルホルムアミド(30.0mL)を入れた。これに続いて、水素化ナトリウム(1.6g、66.7mmol、1.2当量)を5分間添加した。これに、4-ブロモ-2-フルオロピリジン(3.9g、22.2mmol、1.1当量)を20分間で添加した。得られた溶液を0℃で4時間撹拌した。次いで、反応物を、10mLの塩化アンモニウム水溶液を添加することによってクエンチした。得られた溶液を酢酸エチル(50mL×3)で抽出し、有機層を合わせて、真空下で濃縮した。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。これにより、5.2g(62%)のベンジル4-[2-[(4-ブロモピリジン-2-イル)オキシ]エチル]ピペラジン-1-カルボキシレートを黄色の油として得た。 A 100 mL round bottom flask was charged with benzyl 4-(2-hydroxyethyl)piperazine-1-carboxylate (5.3 g, 20.1 mmol, 1.0 equiv.) and N,N-dimethylformamide (30.0 mL). This was followed by the addition of sodium hydride (1.6 g, 66.7 mmol, 1.2 equiv.) over 5 min. To this was added 4-bromo-2-fluoropyridine (3.9 g, 22.2 mmol, 1.1 equiv.) over 20 min. The resulting solution was stirred at 0°C for 4 h. The reaction was then quenched by adding 10 mL of aqueous ammonium chloride solution. The resulting solution was extracted with ethyl acetate (50 mL x 3) and the organic layers were combined and concentrated under vacuum. The residue was applied onto a silica gel column eluting with dichloromethane/methanol (10:1). This yielded 5.2 g (62%) of benzyl 4-[2-[(4-bromopyridin-2-yl)oxy]ethyl]piperazine-1-carboxylate as a yellow oil.
工程3

窒素不活性雰囲気下でパージし、維持した250mLの丸底フラスコに、PhMe(100mL)中のベンジル4-[2-[(4-ブロモピリジン-2-イル)オキシ]エチル]ピペラジン-1-カルボキシレート(2.1g、5.0mmol、1.0当量)、RuphosPd II(775.0mg、1.0mmol、0.2当量)、およびCs2CO3(4.89g、15.0mmol、3.0当量)の溶液を入れた。得られた混合物を、油浴中で、100℃で一晩撹拌した。反応混合物を水(100mL)でクエンチし、酢酸エチル(100mL×2)で抽出した。有機層を合わせ、減圧下で濃縮して、残渣を、100%酢酸エチルで溶出するシリカゲルカラム上に適用した。これにより、670mg(32%)のtert-ブチル8-[2-(2-[4-[(ベンジルオキシ)カルボニル]ピペラジン-1-イル]エトキシ)ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレートを黄色の固体として得た。 A 250 mL round bottom flask purged and maintained under nitrogen inert atmosphere was charged with a solution of benzyl 4-[2-[(4-bromopyridin-2-yl)oxy]ethyl]piperazine-1-carboxylate (2.1 g, 5.0 mmol, 1.0 equiv.), RuphosPd II (775.0 mg, 1.0 mmol, 0.2 equiv.), and Cs2CO3 (4.89 g, 15.0 mmol, 3.0 equiv.) in PhMe (100 mL). The resulting mixture was stirred overnight at 100 °C in an oil bath. The reaction mixture was quenched with water (100 mL) and extracted with ethyl acetate (100 mL x 2). The organic layers were combined and concentrated under reduced pressure, and the residue was applied onto a silica gel column eluting with 100% ethyl acetate. This gave 670 mg (32%) of tert-butyl 8-[2-(2-[4-[(benzyloxy)carbonyl]piperazin-1-yl]ethoxy)pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octane-3-carboxylate as a yellow solid.
工程4
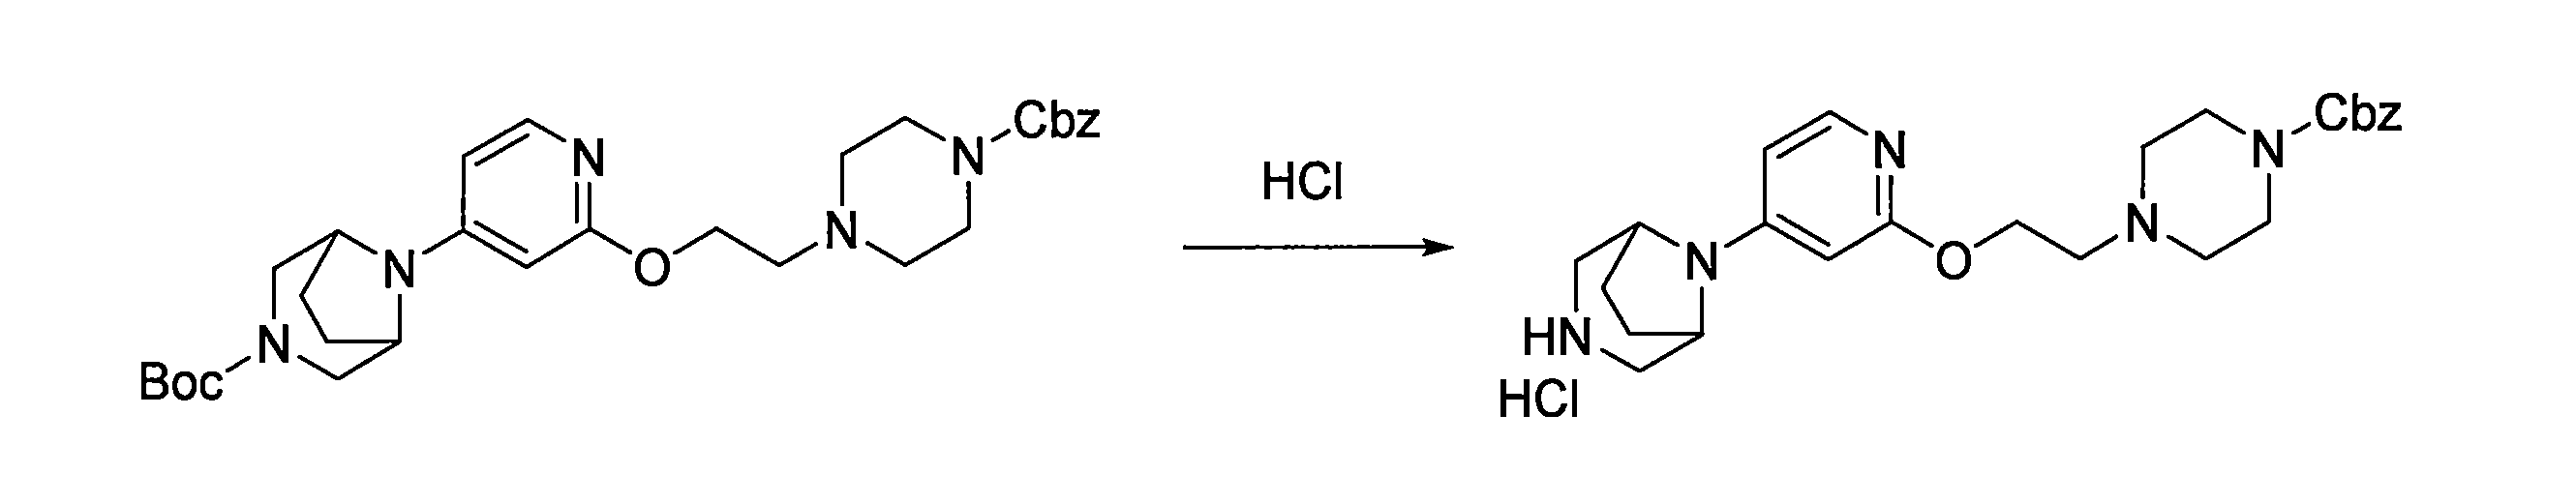
100mLの丸底フラスコに、メタノール(10.0mL)中のtert-ブチル8-[2-(2-[4-[(ベンジルオキシ)カルボニル]ピペラジン-1-イル]エトキシ)ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレート(670mg、1.2mmol、1.0当量)を入れた。次いで、1,4-ジオキサン溶液(4M、10mL)中の塩化水素を添加した。得られた溶液を室温で5時間撹拌した。得られた混合物を真空下で濃縮した。これにより、410mg(69%)のベンジル4-[2-[(4-[3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)オキシ]エチル]ピペラジン-1-カルボキシレート塩酸塩を、黄色の固体として得た。 A 100 mL round bottom flask was charged with tert-butyl 8-[2-(2-[4-[(benzyloxy)carbonyl]piperazin-1-yl]ethoxy)pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octane-3-carboxylate (670 mg, 1.2 mmol, 1.0 equiv.) in methanol (10.0 mL). Hydrogen chloride in 1,4-dioxane solution (4 M, 10 mL) was then added. The resulting solution was stirred at room temperature for 5 hours. The resulting mixture was concentrated under vacuum. This afforded 410 mg (69%) of benzyl 4-[2-[(4-[3,8-diazabicyclo[3.2.1]octane-8-yl]pyridin-2-yl)oxy]ethyl]piperazine-1-carboxylate hydrochloride as a yellow solid.
工程5

10mLの密封チューブに、ベンジル4-[2-[(4-[3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)オキシ]エチル]ピペラジン-1-カルボキシレート塩酸塩(200.0mg、0.4mmol、1.0当量)、4-ブロモ-6-クロロピリダジン-3-アミン(137.0mg、0.7mmol、1.5当量)、メチルスルホキシド(3.0mL)、およびN,N-ジイソプロピルエチルアミン(1mL)を入れた。最終反応混合物を、マイクロ波放射で、130℃で6時間照射した。次いで、反応物を、1mLの水を添加することによってクエンチした。得られた溶液を酢酸エチル(20mL×3)で抽出し、有機層を合わせて、真空下で濃縮した。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。これにより、100mg(42%)のベンジル4-[2-([4-[3-(3-アミノ-6-クロロピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル]オキシ)エチル]ピペラジン-1-カルボキシレートを茶色の固体として得た。 A 10 mL sealed tube was charged with benzyl 4-[2-[(4-[3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl)oxy]ethyl]piperazine-1-carboxylate hydrochloride (200.0 mg, 0.4 mmol, 1.0 equiv.), 4-bromo-6-chloropyridazin-3-amine (137.0 mg, 0.7 mmol, 1.5 equiv.), methyl sulfoxide (3.0 mL), and N,N-diisopropylethylamine (1 mL). The final reaction mixture was irradiated with microwave radiation at 130° C. for 6 h. The reaction was then quenched by adding 1 mL of water. The resulting solution was extracted with ethyl acetate (20 mL×3) and the organic layers were combined and concentrated under vacuum. The residue was applied onto a silica gel column eluting with dichloromethane/methanol (10:1). This gave 100 mg (42%) of benzyl 4-[2-([4-[3-(3-amino-6-chloropyridazin-4-yl)-3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl]oxy)ethyl]piperazine-1-carboxylate as a brown solid.
工程6
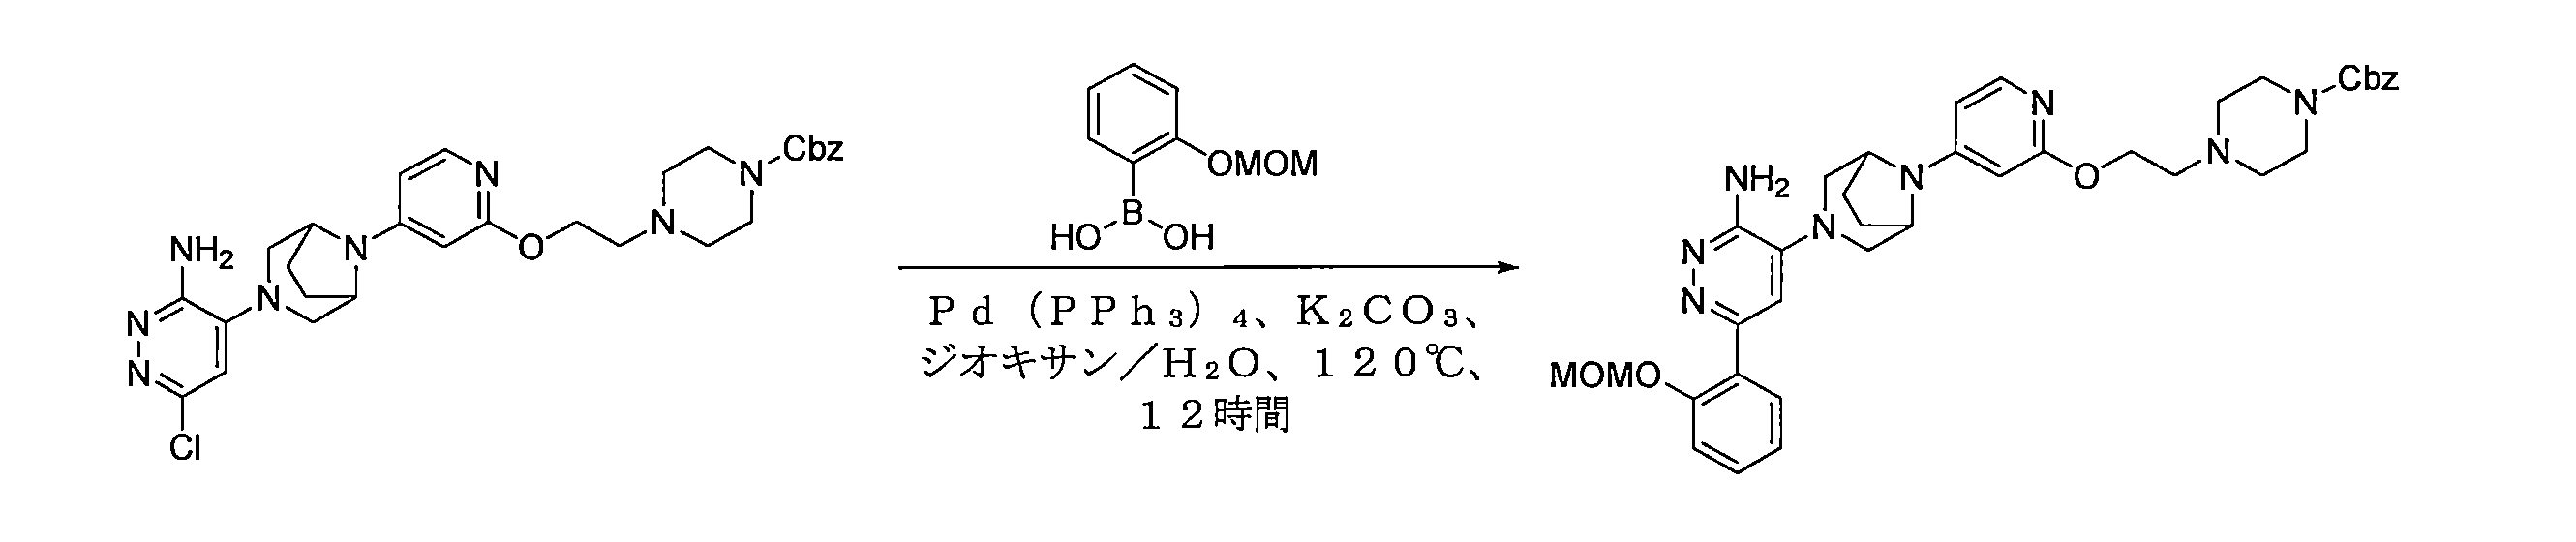
100mLの丸底フラスコに、ベンジル4-[2-([4-[3-(3-アミノ-6-クロロピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル]オキシ)エチル]ピペラジン-1-カルボキシレート(390mg、0.7mmol、1.0当量)、ジオキサン(8.0mL)、水(2.0mL)、炭酸カリウム(279mg、2.0mmol、3.0当量)、[2-(メトキシメトキシ)フェニル]ボロン酸(184.0mg、1.0mmol、1.50当量)、Pd(PPh3)4(78mg、0.07mmol、0.1当量)を入れた。得られた溶液を100℃で2時間撹拌した。得られた溶液を酢酸エチル(20mL×3)で抽出し、有機層を合わせ、真空下で濃縮した。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。これにより、120mg(26%)のベンジル4-(2-[[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]オキシ]エチル)ピペラジン-1-カルボキシレートを茶色の固体として得た。 A 100 mL round bottom flask was charged with benzyl 4-[2-([4-[3-(3-amino-6-chloropyridazin-4-yl)-3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl]oxy)ethyl]piperazine-1-carboxylate (390 mg, 0.7 mmol, 1.0 equiv.), dioxane (8.0 mL), water (2.0 mL), potassium carbonate (279 mg, 2.0 mmol, 3.0 equiv.), [2-(methoxymethoxy)phenyl]boronic acid (184.0 mg, 1.0 mmol, 1.50 equiv.), and Pd(PPh3)4 (78 mg, 0.07 mmol, 0.1 equiv.). The resulting solution was stirred at 100°C for 2 h. The resulting solution was extracted with ethyl acetate (20 mL x 3), and the organic layers were combined and concentrated under vacuum. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (10:1). This gave 120 mg (26%) of benzyl 4-(2-[[4-(3-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl]oxy]ethyl)piperazine-1-carboxylate as a brown solid.
工程7

10mLのイソプロパノール中のベンジル4-(2-[[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]オキシ]エチル)ピペラジン-1-カルボキシレート(200mg、0.29mmol、1.00当量)の溶液に、水酸化パラジウム(100mg、0.71mmol、2.42当量)を、100mLの丸底フラスコ中で、窒素雰囲気下で添加した。次いで、フラスコを真空化し、水素でフラッシュした。得られた溶液を室温で12時間撹拌した。固形分を濾過した。得られた混合物を真空下で濃縮した。これにより、70mg(44%)の6-[2-(メトキシメトキシ)フェニル]-4-(8-[2-[2-(ピペラジン-1-イル)エトキシ]ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-イル)ピリダジン-3-アミンを茶色の油として得た。 To a solution of benzyl 4-(2-[[4-(3-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl]oxy]ethyl)piperazine-1-carboxylate (200 mg, 0.29 mmol, 1.00 equiv.) in 10 mL of isopropanol, palladium hydroxide (100 mg, 0.71 mmol, 2.42 equiv.) was added in a 100 mL round-bottom flask under nitrogen atmosphere. The flask was then evacuated and flushed with hydrogen. The resulting solution was stirred at room temperature for 12 hours. The solids were filtered. The resulting mixture was concentrated under vacuum. This gave 70 mg (44%) of 6-[2-(methoxymethoxy)phenyl]-4-(8-[2-[2-(piperazin-1-yl)ethoxy]pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octan-3-yl)pyridazin-3-amine as a brown oil.
工程8

100mLの丸底フラスコに、4-(テトラメチル-1,3,2-ジオキサボロラン-2-イル)-1H-ピラゾール(1.9g、10.0mmol、1.00当量)、N,N-ジメチルホルムアミド(30mL)、エチル2-ブロモ-3-メチルブタノエート(2.1g、10.0mmol、1.0当量)、および炭酸セシウム(10.0g、30.7mmol、3.0当量)を入れた。得られた溶液を90℃で12時間撹拌した。次いで、反応物を、10mLの水を添加することによってクエンチした。得られた溶液を酢酸エチル(20mL×3)で抽出し、有機層を合わせて、真空下で濃縮した。残渣を、酢酸エチル/石油エーテル(1:6)で溶出するシリカゲルカラム上に適用した。これにより、3.3g(93%)のエチル3-メチル-2-[4-(テトラメチル-1,3,2-ジオキサボロラン-2-イル)-1H-ピラゾール-1-イル]ブタノエートを白色の固体として得た。 A 100 mL round bottom flask was charged with 4-(tetramethyl-1,3,2-dioxaborolan-2-yl)-1H-pyrazole (1.9 g, 10.0 mmol, 1.00 equiv.), N,N-dimethylformamide (30 mL), ethyl 2-bromo-3-methylbutanoate (2.1 g, 10.0 mmol, 1.0 equiv.), and cesium carbonate (10.0 g, 30.7 mmol, 3.0 equiv.). The resulting solution was stirred at 90° C. for 12 h. The reaction was then quenched by adding 10 mL of water. The resulting solution was extracted with ethyl acetate (20 mL×3) and the organic layers were combined and concentrated under vacuum. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (1:6). This yielded 3.3 g (93%) of ethyl 3-methyl-2-[4-(tetramethyl-1,3,2-dioxaborolan-2-yl)-1H-pyrazol-1-yl]butanoate as a white solid.
工程9

250mLの丸底フラスコに、エチル3-メチル-2-[4-(テトラメチル-1,3,2-ジオキサボロラン-2-イル)-1H-ピラゾール-1-イル]ブタノエート(3.0g、9.3mmol、1.0当量)、テトラヒドロフラン(30mL)、水(30mL)、過ホウ酸ナトリウム(3.5g、23.3mmol、2.5当量)を入れた。得られた溶液を室温で12時間撹拌した。得られた混合物を真空下で濃縮した。得られた溶液を酢酸エチル(20mL×3)で抽出し、有機層を合わせて、真空下で濃縮した。残渣を、酢酸エチル/石油エーテル(1:1)で溶出するシリカゲルカラム上に適用した。これにより、1740mg(88%)のエチル2-(4-ヒドロキシ-1H-ピラゾール-1-イル)-3-メチルブタノエートを白色の固体として得た。 A 250 mL round bottom flask was charged with ethyl 3-methyl-2-[4-(tetramethyl-1,3,2-dioxaborolan-2-yl)-1H-pyrazol-1-yl]butanoate (3.0 g, 9.3 mmol, 1.0 equiv.), tetrahydrofuran (30 mL), water (30 mL), and sodium perborate (3.5 g, 23.3 mmol, 2.5 equiv.). The resulting solution was stirred at room temperature for 12 hours. The resulting mixture was concentrated under vacuum. The resulting solution was extracted with ethyl acetate (20 mL x 3), and the organic layers were combined and concentrated under vacuum. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (1:1). This afforded 1740 mg (88%) of ethyl 2-(4-hydroxy-1H-pyrazol-1-yl)-3-methylbutanoate as a white solid.
工程10
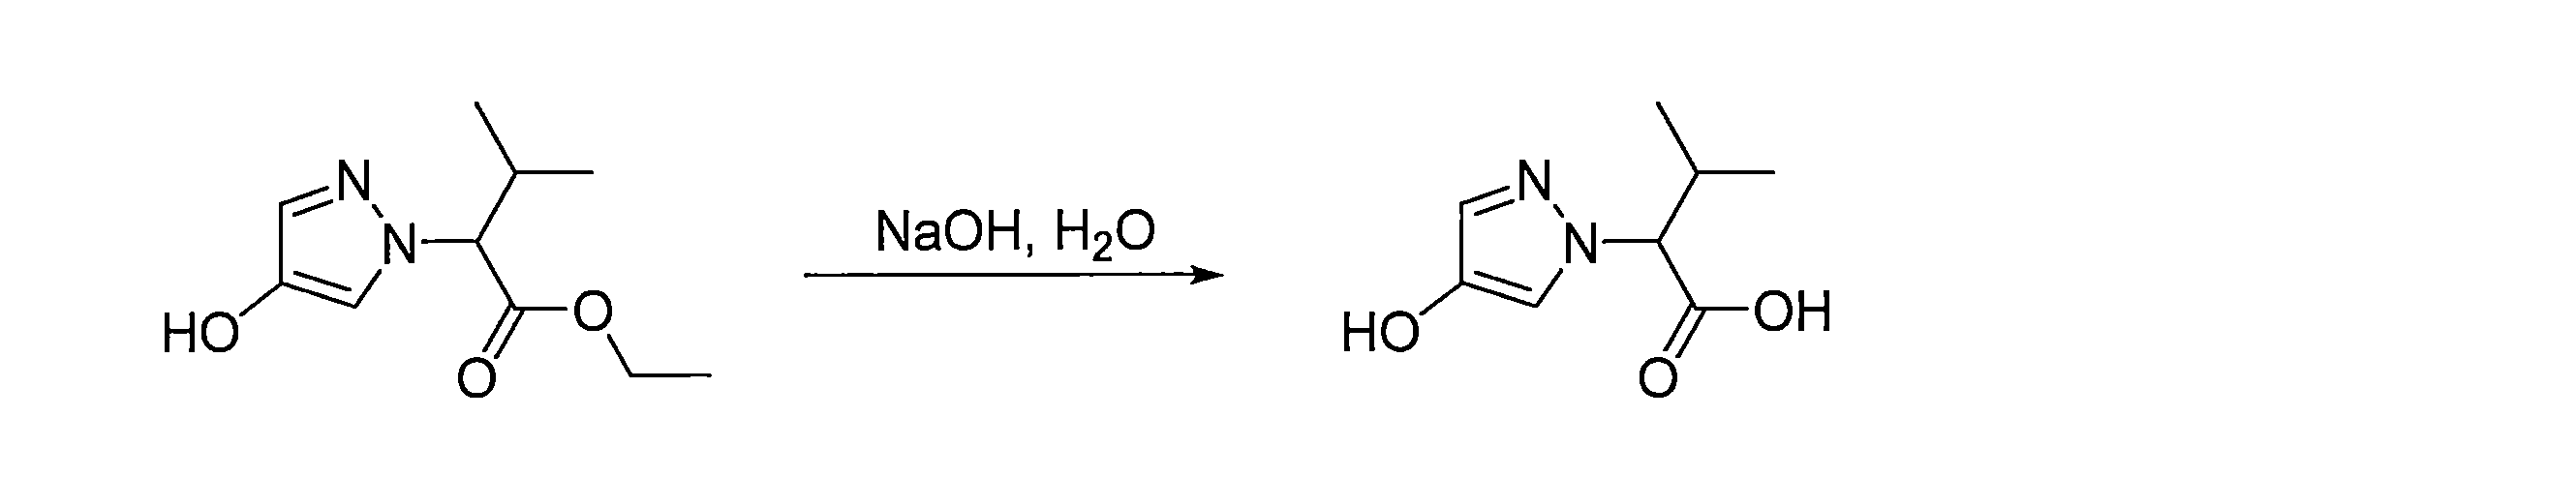
50mLの丸底フラスコに、エチル2-(4-ヒドロキシ-1H-ピラゾール-1-イル)-3-メチルブタノエート(3.5g、16.5mmol、1.0当量)、メタノール(10mL)、水(10mL)、および水酸化ナトリウム(2.7g、67.5mmol、4.0当量)を入れた。得られた溶液を室温で12時間撹拌した。溶液のpH値を、2M HClで2に調整した。以下の条件の分取HPLCによって、粗生成物を精製した:カラム、XBridge Shield RP18 OBDカラム、5um、19*150mm;移動相、水(0.1%FA)およびアセトニトリル(0.0%アセトニトリル、2分間で最大4.0%、8分間で最大20.0%);検出器、UV254nm。これにより、1.5g(49%)の2-(4-ヒドロキシ-1H-ピラゾール-1-イル)-3-メチルブタン酸を白色の油として得た。 A 50 mL round bottom flask was charged with ethyl 2-(4-hydroxy-1H-pyrazol-1-yl)-3-methylbutanoate (3.5 g, 16.5 mmol, 1.0 equiv.), methanol (10 mL), water (10 mL), and sodium hydroxide (2.7 g, 67.5 mmol, 4.0 equiv.). The resulting solution was stirred at room temperature for 12 hours. The pH value of the solution was adjusted to 2 with 2M HCl. The crude product was purified by preparative HPLC with the following conditions: column, XBridge Shield RP18 OBD column, 5 um, 19*150 mm; mobile phase, water (0.1% FA) and acetonitrile (0.0% acetonitrile, max. 4.0% in 2 min, max. 20.0% in 8 min); detector, UV 254 nm. This gave 1.5 g (49%) of 2-(4-hydroxy-1H-pyrazol-1-yl)-3-methylbutanoic acid as a white oil.
工程11

100mLの丸底フラスコに、2-(4-ヒドロキシ-1H-ピラゾール-1-イル)-3-メチルブタン酸(500mg、2.7mmol、1.0当量)、N,N-ジメチルホルムアミド(20mL)、(2S,4R)-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド塩酸塩(900mg、2.5mmol、0.9当量)、DIEA(1.1g、8.5mmol、3.0当量)、T3P(2.6g、1.5当量)を入れた。得られた溶液を室温で1時間撹拌した。次いで、反応物を、5mLの水を添加することによってクエンチした。得られた溶液を酢酸エチル(30mL×3)で抽出し、有機層を合わせて、真空下で濃縮した。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。これにより、800mg(59%)の(2S,4R)-4-ヒドロキシ-1-[2-(4-ヒドロキシ-1H-ピラゾール-1-イル)-3-メチルブタノイル]-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミドを黄色の固体として得た。 A 100 mL round bottom flask was charged with 2-(4-hydroxy-1H-pyrazol-1-yl)-3-methylbutanoic acid (500 mg, 2.7 mmol, 1.0 equiv.), N,N-dimethylformamide (20 mL), (2S,4R)-4-hydroxy-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide hydrochloride (900 mg, 2.5 mmol, 0.9 equiv.), DIEA (1.1 g, 8.5 mmol, 3.0 equiv.), and T3P (2.6 g, 1.5 equiv.). The resulting solution was stirred at room temperature for 1 h. The reaction was then quenched by adding 5 mL of water. The resulting solution was extracted with ethyl acetate (30 mL x 3) and the organic layers were combined and concentrated under vacuum. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (10:1). This gave 800 mg (59%) of (2S,4R)-4-hydroxy-1-[2-(4-hydroxy-1H-pyrazol-1-yl)-3-methylbutanoyl]-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide as a yellow solid.
工程12

5mLの密封チューブに、(2S,4R)-4-ヒドロキシ-1-[2-(4-ヒドロキシ-1H-ピラゾール-1-イル)-3-メチルブタノイル]-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(300mg、0.60mmol、1.0当量)、N,N-ジメチルホルムアミド(5.0mL)、1,2-ジブロモエタン(222mg、1.2mmol、2.0当量)、炭酸カリウム(250mg、1.8mmol、3.0当量)を入れた。得られた溶液を70℃で12時間撹拌した。次いで、反応物を、1mLの水を添加することによってクエンチした。以下の条件の分取HPLCによって、粗生成物を精製した:カラム、XBridge Shield RP18 OBDカラム、5um、19*150mm;移動相、水(10mmol/Lの重炭酸アンモニウム)およびアセトニトリル(35.0%アセトニトリル、8分間で最大47.0%);検出器、UV254nm。これにより、120mg(33%)の(2S,4R)-1-[2-[4-(2-ブロモエトキシ)-1H-ピラゾール-1-イル]-3-メチルブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミドを黄色の固体として得た。 A 5 mL sealed tube was charged with (2S,4R)-4-hydroxy-1-[2-(4-hydroxy-1H-pyrazol-1-yl)-3-methylbutanoyl]-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (300 mg, 0.60 mmol, 1.0 equiv.), N,N-dimethylformamide (5.0 mL), 1,2-dibromoethane (222 mg, 1.2 mmol, 2.0 equiv.), and potassium carbonate (250 mg, 1.8 mmol, 3.0 equiv.). The resulting solution was stirred at 70° C. for 12 h. The reaction was then quenched by adding 1 mL of water. The crude product was purified by preparative HPLC with the following conditions: column, XBridge Shield RP18 OBD column, 5um, 19*150mm; mobile phase, water (10mmol/L ammonium bicarbonate) and acetonitrile (35.0% acetonitrile, max 47.0% in 8 min); detector, UV 254nm. This gave 120mg (33%) of (2S,4R)-1-[2-[4-(2-bromoethoxy)-1H-pyrazol-1-yl]-3-methylbutanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide as a yellow solid.
工程13

100mLの丸底フラスコに、6-[2-(メトキシメトキシ)フェニル]-4-(8-[2-[2-(ピペラジン-1-イル)エトキシ]ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-イル)ピリダジン-3-アミン(54.6mg、0.1mmol、1.0当量)、アセトニトリル(10.0mL)、炭酸カリウム(41mg、0.30mmol、3.0当量)、ヨウ化ナトリウム(18mg、0.1mmol、1.2当量)、(2S,4R)-1-2-[4-(2-ブロモエトキシ)-1H-ピラゾール-1-イル]-3-メチルブタノイル-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5yl)フェニル]エチル]ピロリジン-2-カルボキサミド(60mg、0.1mmol、1.0当量)を入れた。得られた溶液を70℃で12時間撹拌した。次いで、反応物を、3mLの水を添加することによってクエンチした。得られた溶液を酢酸エチル(30mL×3)で抽出し、有機層を合わせて、真空下で濃縮した。残渣を、ジクロロメタン/メタノール(10/1)で溶出するシリカゲルカラム上に適用した。これにより、30mg(29%)の(2S,4R)-1-(2-[4-[2-(4-[2-[(4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)オキシ]エチル]ピペラジン-1-イル)エトキシ]-1H-ピラゾール-1-イル]-3-メチルブタノイル)-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミドを淡い茶色の固体として得た。 In a 100 mL round bottom flask, add 6-[2-(methoxymethoxy)phenyl]-4-(8-[2-[2-(piperazin-1-yl)ethoxy]pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octan-3-yl)pyridazin-3-amine (54.6 mg, 0.1 mmol, 1.0 equiv.), acetonitrile (10.0 mL), potassium carbonate (41 mg, 0.30 mmol, 3.0 eq.), sodium iodide (18 mg, 0.1 mmol, 1.2 eq.), (2S,4R)-1-2-[4-(2-bromoethoxy)-1H-pyrazol-1-yl]-3-methylbutanoyl-4-hydroxy-N-[(1S)-1-[4-(4-methyl-1,3-thiazole-5yl)phenyl]ethyl]pyrrolidine-2-carboxamide (60 mg, 0.1 mmol, 1.0 eq.). The resulting solution was stirred at 70° C. for 12 hours. The reaction was then quenched by adding 3 mL of water. The resulting solution was extracted with ethyl acetate (30 mL×3) and the organic layers were combined and concentrated under vacuum. The residue was applied onto a silica gel column eluting with dichloromethane/methanol (10/1). This gave 30 mg (29%) of (2S,4R)-1-(2-[4-[2-(4-[2-[(4-[3-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl)oxy]ethyl]piperazin-1-yl)ethoxy]-1H-pyrazol-1-yl]-3-methylbutanoyl)-4-hydroxy-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide as a light brown solid.
工程14

25mLの丸底フラスコに、(2S,4R)-1-[2-(4-[2-[4-(2-[[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]オキシ]エチル)ピペラジン-1-イル]エトキシ]-1H-ピラゾール-1-イル)-3-メチルブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(40mg、0.04mmol、1.0当量)、メタノール(8mL)、塩化水素(150mL)を入れた。得られた溶液を室温で6時間撹拌した。次いで、反応物を、1mLの水を添加することによってクエンチした。粗生成物(5mL)を、以下の条件を用いて、分取HPLCによって精製した。カラム、XBridge Shield RP18 OBDカラム、5um、19*150mm;移動相、水(0.1%ギ酸)およびアセトニトリル(13.0%アセトニトリル、8分間で最大27.0%);検出器、UV254nm。これにより、30mg(78%)の(2S,4R)-1-(2-[4-[2-(4-[2-[(4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)オキシ]エチル]ピペラジン-1-イル)エトキシ]-1H-ピラゾール-1-イル]-3-メチルブタノイル)-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミドを白色の固体として得た。 A 25 mL round bottom flask was charged with (2S,4R)-1-[2-(4-[2-[4-(2-[[4-(3-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl]oxy]ethyl)piperazin-1-yl]ethoxy]-1H-pyrazol-1-yl)-3-methylbutanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (40 mg, 0.04 mmol, 1.0 equiv.), methanol (8 mL), and hydrogen chloride (150 mL). The resulting solution was stirred at room temperature for 6 hours. The reaction was then quenched by adding 1 mL of water. The crude product (5 mL) was purified by preparative HPLC using the following conditions: column, XBridge Shield RP18 OBD column, 5 um, 19*150 mm; mobile phase, water (0.1% formic acid) and acetonitrile (13.0% acetonitrile, max 27.0% in 8 min); detector, UV 254 nm. This gave 30 mg (78%) of (2S,4R)-1-(2-[4-[2-(4-[2-[(4-[3-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl)oxy]ethyl]piperazin-1-yl)ethoxy]-1H-pyrazol-1-yl]-3-methylbutanoyl)-4-hydroxy-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide as a white solid.
(2S,4R)-1-(2-[4-[2-(4-[2-[(4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)オキシ]エチル]ピペラジン-1-イル)エトキシ]-1H-ピラゾール-1-イル]-3-メチルブタノイル)-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミドを、以下の条件:カラム、Chiralpak IC、2*25cm、5um;移動相、DCMおよびメタノール(11分で50.0%メタノールを保持);検出器、UV220/254nmを用いて、キラル分取HPLCによって精製した。これにより、5.6mgの(2S,4R)-1-[(2R)-2-[4-[2-(4-[2-[(4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)オキシ]エチル]ピペラジン-1-イル)エトキシ]-1H-ピラゾール-1-イル]-3-メチルブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミドを白色の固体[1H NMR(300MHz、メタノール-d4)δ8.89(s、1H)、7.85~7.72(m、2H)、7.56~7.36(m、6H)、7.31~7.16(m、2H)、6.97~6.85(m、2H)、6.58(dd、J=6.2、2.2Hz、1H)、6.24(d、J=2.1Hz、1H)、5.05(q、J=6.9Hz、1H)、4.67(d、J=10.6Hz、1H)、4.60~4.47(m、3H)、4.46(s、1H)、4.39(t、J=5.4Hz、2H)、4.06(t、J=5.3Hz、2H)、3.87(dd、J=10.9、4.2Hz、1H)、3.76(s、1H)、3.73~3.55(m、2H)、3.19~2.99(m、2H)、2.92~2.69(m、12H)、2.50(s、4H)、2.30~2.14(m、5H)、1.98(ddd、J=13.2、8.7、4.6Hz、1H)、1.54(d、J=7.0Hz、3H)、1.41~1.26(m、4H)、1.20(t、J=7.0Hz、2H)、1.08(d、J=6.5Hz、3H)、0.93(d、J=4.1Hz、1H)]として、および8.1mgの(2S,4R)-1-[(2S)-2-[4-[2-(4-[2-[(4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)オキシ]エチル]ピペラジン-1-イル)エトキシ]-1H-ピラゾール-1-イル]-3-メチルブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミドを白色の固体[1H NMR(300MHz、メタノール-d4)δ8.88(d、J=13.8Hz、1H)、7.85~7.69(m、2H)、7.59~7.30(m、6H)、7.32~7.18(m、2H)、6.97~6.84(m、2H)、6.57(dd、J=6.1、2.1Hz、1H)、6.23(t、J=2.7Hz、1H)、4.99(q、J=7.0Hz、1H)、4.78(d、J=8.8Hz、1H)、4.60(t、J=8.2Hz、1H)、4.53(s、2H)、4.37(dd、J=6.8、4.1Hz、3H)、4.04(t、J=5.3Hz、2H)、3.64(p、J=6.7Hz、3H)、3.40~3.24(m、1H)、3.11(dd、J=15.8、4.6Hz、2H)、2.88~2.64(m、12H)、2.49(d、J=13.8Hz、4H)、2.34~2.10(m、6H)、2.06(s、1H)、1.95(td、J=8.5、4.2Hz、1H)、1.50(d、J=7.0Hz、3H)、1.33(td、J=7.8、7.4、4.4Hz、5H)、1.20(t、J=7.0Hz、3H)、1.08(d、J=6.7Hz、3H)、0.93(q、J=6.8Hz、1H)、0.74(dd、J=19.3、6.7Hz、3H)]として得た。 (2S,4R)-1-(2-[4-[2-(4-[2-[(4-[3-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl)oxy]ethyl]piperazin-1-yl)ethoxy]-1H-pyrazol-1-yl]-3-methylbutanoyl)-4-hydroxy-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide was purified by chiral preparative HPLC using the following conditions: column, Chiralpak IC, 2*25cm, 5um; mobile phase, DCM and methanol (50.0% methanol held at 11 min); detector, UV 220/254nm. This gave 5.6 mg of (2S,4R)-1-[(2R)-2-[4-[2-(4-[2-[(4-[3-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl)oxy]ethyl]piperazin-1-yl)ethoxy]-1H-pyrazol-1-yl]-3-methylbutanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide as a white solid [1H NMR (300 MHz, methanol-d4) δ 8.89 (s, 1H), 7.85-7.72 (m, 2H), 7.56-7.36 (m, 6H), 7.31-7.16 (m, 2H), 6.97-6.85 (m, 2H), 6.58 (dd, J = 6.2, 2.2 Hz, 1H), 6.24 (d, J = 2.1 Hz, 1H), 5.05 (q, J = 6.9 Hz, 1H), 4.67 (d, J = 10.6 Hz, 1H), 4.6 0-4.47 (m, 3H), 4.46 (s, 1H), 4.39 (t, J = 5.4 Hz, 2H), 4.06 (t, J = 5.3 Hz, 2H), 3.87 (dd, J = 10.9, 4.2 Hz, 1H), 3.76 (s, 1H), 3.73-3.55 (m, 2H), 3.19-2.99 (m, 2H), 2.92-2.69 (m, 12H), 2.50 (s, 4H), 2.30-2.14 (m, 5H), 1.98 (ddd , J = 13.2, 8.7, 4.6 Hz, 1H), 1.54 (d, J = 7.0 Hz, 3H), 1.41-1.26 (m, 4H), 1.20 (t, J = 7.0 Hz, 2H), 1.08 (d, J = 6.5 Hz, 3H), 0.93 (d, J = 4.1 Hz, 1H)], and 8.1 mg of (2S,4R)-1-[(2S)-2-[4-[2-(4-[2-[(4-[3-[3-amino-6-(2-hydroxyphenyl)-2-propanediol, 1H], 4-hydroxy-2-(2-hydroxyphenyl)-1-(2-propanediol, 1H)]. [1H-pyrazol-1-yl]-3-methylbutanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide was obtained as a white solid [1H NMR (300 MHz, methanol-d4) δ 8.88 (d, J = 13.8 Hz, 1H), 7.85-7.69 (m, 2H), 7.59-7.30 (m, 6H), 7.32-7.18 (m, 2H), 6.97-6.84 (m, 2H), 6.57 (dd, J = 6.1, 2.1 Hz, 1H), 6.23 (t, J = 2.7 Hz, 1H), 4.99 (q, J = 7.0 Hz, 1H), 4.78 (d, J = 8.8 Hz, 1H), 4.60 (t, J = 8.2 Hz, 1H), 4.53 (s, 2H), 4.37 (dd, J = 6.8, 4.1 Hz, 3H), 4.04 (t, J = 5.3 Hz, 2H), 3.64 (p, J = 6.7 Hz, 3H), 3.40-3.24 (m, 1H), 3.11 (dd, J = 15.8, 4.6 Hz, 2H), 2.88-2.64 (m, 12H), 2.49 (d, J = 13.8 Hz, 4H), 2.34-2.10 (m, 6H), 2.06 (s, 1H), 1.95 (td, J = 8.5, 4.2 Hz, 1H), 1.50 (d, J = 7.0 Hz, 3H), 1.33 (td, J = 7.8, 7.4, 4.4 Hz, 5H), 1.20 (t, J = 7.0 Hz, 3H), 1.08 (d, J = 6.7 Hz, 3H), 0.93 (q, J = 6.8 Hz, 1H), 0.74 (dd, J = 19.3, 6.7 Hz, 3H)].
例示的な化合物43および例示的な化合物44の例示的な合成
工程1

250mLの丸底フラスコに、NMP(120mL)中の2-ブロモ-4-フルオロピリジン(7.8g、44.32mmol、1.05当量)、tert-ブチル3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレート(9g、42.4mmol、1.00当量)、DIEA(25g、193mmol、4.00当量)の溶液を入れた。得られた溶液を、150℃で3時間撹拌した。反応混合物を冷却した。次いで、反応物を、水(200mL)を添加することによってクエンチした。得られた溶液を酢酸エチル(50mL×3)で抽出し、有機層を合わせた。得られた混合物を、水(50mL×1)およびブライン(50mL×1)で洗浄した。混合物を、無水硫酸ナトリウム上で乾燥させ、真空下で濃縮した。残渣を、酢酸エチル/石油エーテル(1:2)で溶出するシリカゲルカラム上に適用した。これにより、11.7g(75%)のtert-ブチル8-(2-ブロモピリジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレートを黄色の固体として得た。 A 250 mL round bottom flask was charged with a solution of 2-bromo-4-fluoropyridine (7.8 g, 44.32 mmol, 1.05 equiv.), tert-butyl 3,8-diazabicyclo[3.2.1]octane-3-carboxylate (9 g, 42.4 mmol, 1.00 equiv.), DIEA (25 g, 193 mmol, 4.00 equiv.) in NMP (120 mL). The resulting solution was stirred at 150 °C for 3 h. The reaction mixture was cooled. The reaction was then quenched by adding water (200 mL). The resulting solution was extracted with ethyl acetate (50 mL x 3) and the organic layers were combined. The resulting mixture was washed with water (50 mL x 1) and brine (50 mL x 1). The mixture was dried over anhydrous sodium sulfate and concentrated under vacuum. The residue was applied onto a silica gel column eluting with ethyl acetate/petroleum ether (1:2). This yielded 11.7 g (75%) of tert-butyl 8-(2-bromopyridin-4-yl)-3,8-diazabicyclo[3.2.1]octane-3-carboxylate as a yellow solid.
工程2

250mLの丸底フラスコに、[(プロパ-2-イン-1-イルオキシ)メチル]ベンゼン(10g、68.4mmol、1.00当量)、ピナコールボラン(10mL)、ZrCp2HCl(2g)、トリエチルアミン(850mg、8.40mmol、0.12当量)の混合物を入れた。得られた溶液を、油浴中で、65℃で16時間撹拌した。得られた混合物を水(30mL)でクエンチし、次いで、酢酸エチル(50mL×3)で抽出して、有機層を合わせた。得られた混合物をブライン(50mL×1)で洗浄した。得られた混合物を真空下で濃縮した。残渣を、酢酸エチル/石油エーテル(1:1)で溶出するシリカゲルカラム上に適用した。これにより、12.5g(67%)の2-[(1E)-3-(ベンジルオキシ)プロパ-1-エン-1-イル]-4,4,5,5-テトラメチル-1,3,2-ジオキサボロランを黄色の固体として得た。 A mixture of [(prop-2-yn-1-yloxy)methyl]benzene (10 g, 68.4 mmol, 1.00 equiv.), pinacolborane (10 mL), ZrCp2HCl (2 g), and triethylamine (850 mg, 8.40 mmol, 0.12 equiv.) was placed in a 250 mL round-bottom flask. The resulting solution was stirred in an oil bath at 65°C for 16 h. The resulting mixture was quenched with water (30 mL) and then extracted with ethyl acetate (50 mL x 3) and the organic layers were combined. The resulting mixture was washed with brine (50 mL x 1). The resulting mixture was concentrated under vacuum. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (1:1). This yielded 12.5 g (67%) of 2-[(1E)-3-(benzyloxy)prop-1-en-1-yl]-4,4,5,5-tetramethyl-1,3,2-dioxaborolane as a yellow solid.
工程3
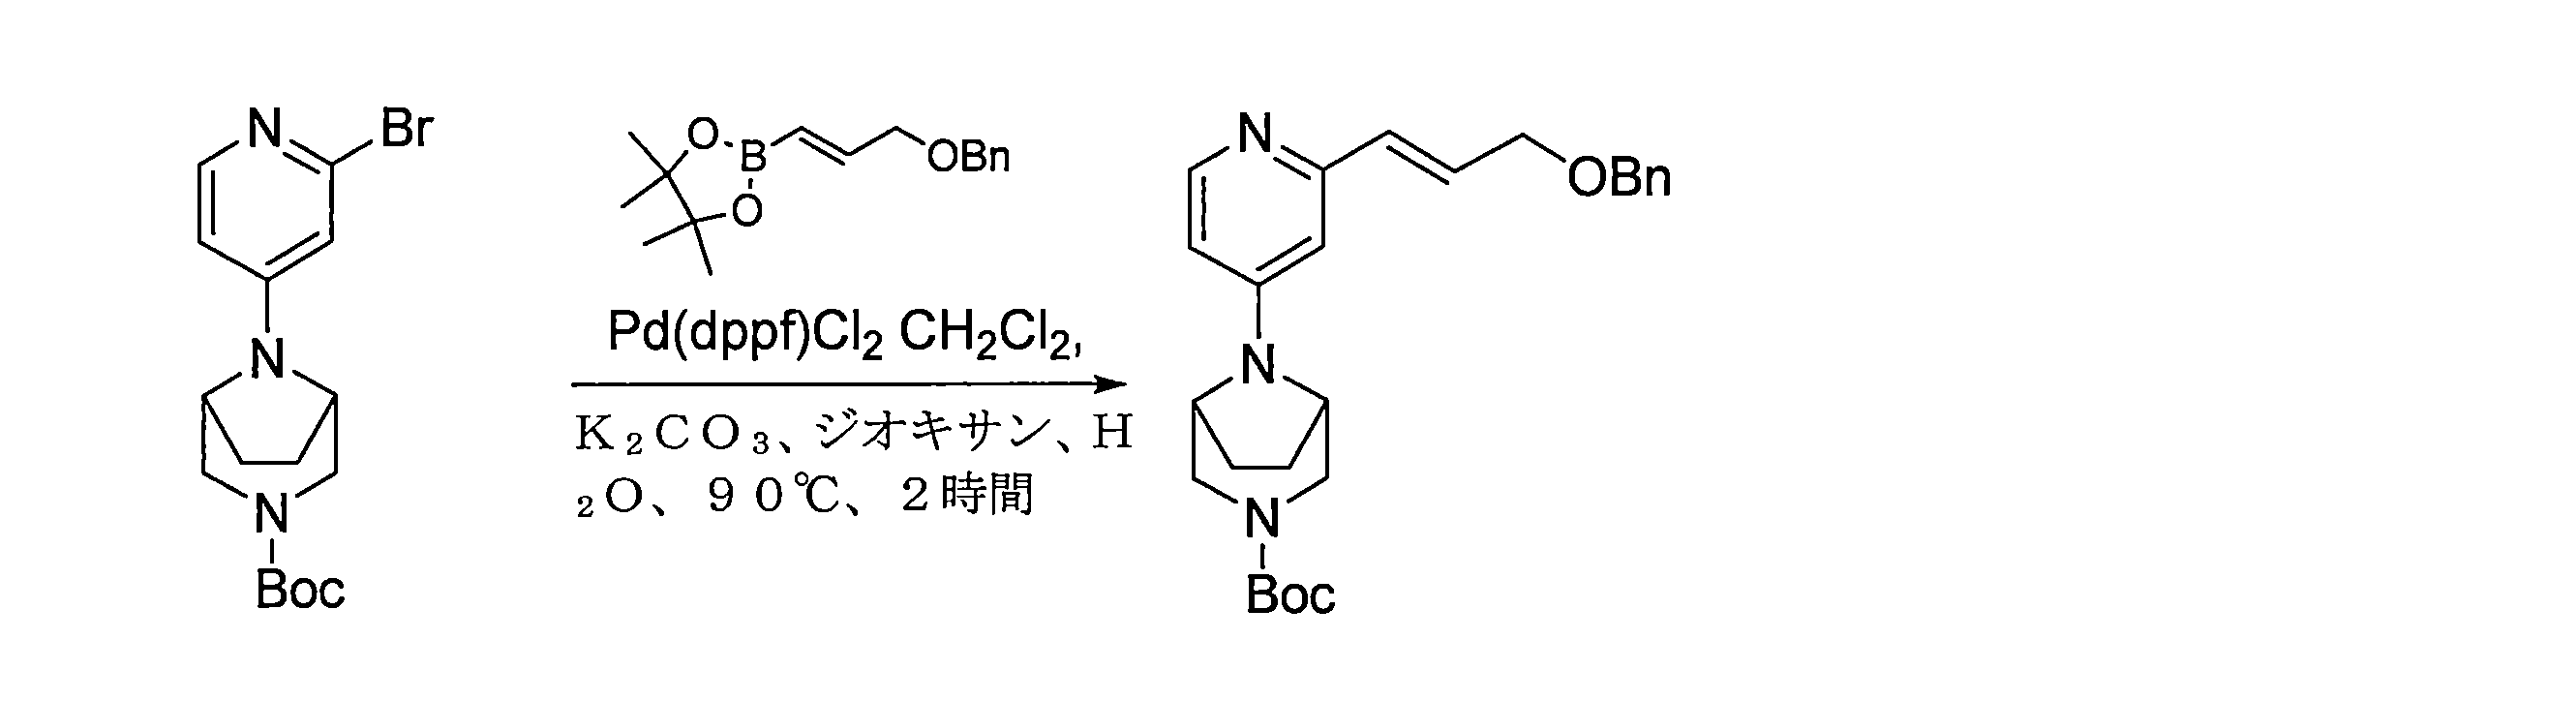
窒素不活性雰囲気下でパージし、維持した250mLの丸底フラスコに、ジオキサン/H2O(80/20mL)中のtert-ブチル8-(2-ブロモピリジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレート(4.0g、10.9mmol、1.00当量)、2-[(1E)-3-(ベンジルオキシ)プロパ-1-エン-1-イル]-4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン(4.48g、16.3mmol、1.50当量)、Pd(dppf)2Cl2ジクロロメタン(890mg、1.09mmol、0.10当量)、および炭酸カリウム(4.5g、32.6mmol、3.00当量)を入れた。得られた溶液を、90℃で2時間撹拌した。反応混合物を室温に冷却した。得られた混合物を真空下で濃縮した。残渣を、酢酸エチル/石油エーテル(1:1)で溶出するシリカゲルカラム上に適用した。これにより、4.55g(96%)のtert-ブチル8-[2-[(1E)-3-(ベンジルオキシ)プロパ-1-エン-1-イル]ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレートを黄色の固体として得た。 A 250 mL round bottom flask purged and maintained under a nitrogen inert atmosphere was charged with tert-butyl 8-(2-bromopyridin-4-yl)-3,8-diazabicyclo[3.2.1]octane-3-carboxylate (4.0 g, 10.9 mmol, 1.00 equiv.), 2-[(1E)-3-(benzyloxy)prop-1-en-1-yl]-4,4,5,5-tetramethyl-1,3,2-dioxaborolane (4.48 g, 16.3 mmol, 1.50 equiv.), Pd(dppf)2Cl2 dichloromethane (890 mg, 1.09 mmol, 0.10 equiv.), and potassium carbonate (4.5 g, 32.6 mmol, 3.00 equiv.) in dioxane/H2O (80/20 mL). The resulting solution was stirred at 90° C. for 2 h. The reaction mixture was cooled to room temperature. The resulting mixture was concentrated under vacuum. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (1:1). This gave 4.55 g (96%) of tert-butyl 8-[2-[(1E)-3-(benzyloxy)prop-1-en-1-yl]pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octane-3-carboxylate as a yellow solid.
工程4

窒素不活性雰囲気下でパージし、維持した250mLの丸底フラスコに、メタノール/HOAc(200/0.5mL)中のtert-ブチル8-[2-[(1E)-3-(ベンジルオキシ)プロパ-1-エン-1-イル]ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレート(12.9g、29.6mmol、1.00当量)の溶液を入れた。炭素上のパラジウム(6g)を窒素雰囲気下で添加した。次いで、フラスコを真空化し、水素でフラッシュした。反応混合物を、水素バルーンを使用して水素雰囲気下で、50℃で20時間水素化した。固形分をセライトパッドを通して濾過し、濾液を減圧下で濃縮した。これにより、10.2g(99%)のtert-ブチル8-[2-(3-ヒドロキシプロピル)ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレートを黄色の油として得た。 A 250 mL round bottom flask purged and maintained under a nitrogen inert atmosphere was charged with a solution of tert-butyl 8-[2-[(1E)-3-(benzyloxy)prop-1-en-1-yl]pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octane-3-carboxylate (12.9 g, 29.6 mmol, 1.00 equiv.) in methanol/HOAc (200/0.5 mL). Palladium on carbon (6 g) was added under a nitrogen atmosphere. The flask was then evacuated and flushed with hydrogen. The reaction mixture was hydrogenated under a hydrogen atmosphere using a hydrogen balloon at 50° C. for 20 hours. The solids were filtered through a celite pad and the filtrate was concentrated under reduced pressure. This yielded 10.2 g (99%) of tert-butyl 8-[2-(3-hydroxypropyl)pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octane-3-carboxylate as a yellow oil.
工程5

100mLの丸底フラスコに、ジクロロメタン(30mL)中のtert-ブチル8-[2-(3-ヒドロキシプロピル)ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレート(700mg、2.01mmol、1.00当量)を入れ、デスマーチンペリオジナン試薬(1.7g、2.00当量)を0~5℃で添加した。得られた溶液を室温で6時間撹拌した。得られた混合物を15℃において真空下で濃縮した。残渣を、ジクロロメタン/メタノール(5:1)で溶出するシリカゲルカラム上に適用した。これにより、500mg(72%)のtert-ブチル8-[2-(3-オキソプロピル)ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレートを白色の固体として得た。 A 100 mL round bottom flask was charged with tert-butyl 8-[2-(3-hydroxypropyl)pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octane-3-carboxylate (700 mg, 2.01 mmol, 1.00 equiv.) in dichloromethane (30 mL) and Dess-Martin periodinane reagent (1.7 g, 2.00 equiv.) was added at 0-5°C. The resulting solution was stirred at room temperature for 6 h. The resulting mixture was concentrated under vacuum at 15°C. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (5:1). This gave 500 mg (72%) of tert-butyl 8-[2-(3-oxopropyl)pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octane-3-carboxylate as a white solid.
工程6

50mLの丸底フラスコに、ジクロロメタン(20mL)中のtert-ブチル8-[2-(3-オキソプロピル)ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレート(1.4g、4.05mmol、1.00当量)、ベンジルピペラジン-1-カルボキシレート(1.07g、4.86mmol、1.20当量)を入れた。これを20分間撹拌し、NaBH(OAc)3(2.58g、12.2mmol、3.00当量)を0℃で添加した。得られた溶液を室温で4時間撹拌した。次いで、反応物を、水/氷(50mL)を添加することによってクエンチした。得られた溶液を、ジクロロメタン(50mL×3)で抽出し、有機層を合わせて、無水硫酸ナトリウムで乾燥させた。残渣を、ジクロロメタン/メタノール(10/1)で溶出するシリカゲルカラム上に適用した。これにより、2.19g(98%)のtert-ブチル8-[2-(3-[4-[(ベンジルオキシ)カルボニル]ピペラジン-1-イル]プロピル)ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレートを淡い黄色の油として得た。 A 50 mL round bottom flask was charged with tert-butyl 8-[2-(3-oxopropyl)pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octane-3-carboxylate (1.4 g, 4.05 mmol, 1.00 equiv.), benzyl piperazine-1-carboxylate (1.07 g, 4.86 mmol, 1.20 equiv.) in dichloromethane (20 mL). This was stirred for 20 minutes and NaBH(OAc)3 (2.58 g, 12.2 mmol, 3.00 equiv.) was added at 0°C. The resulting solution was stirred at room temperature for 4 hours. The reaction was then quenched by adding water/ice (50 mL). The resulting solution was extracted with dichloromethane (50 mL x 3) and the organic layers were combined and dried over anhydrous sodium sulfate. The residue was applied onto a silica gel column eluting with dichloromethane/methanol (10/1). This gave 2.19 g (98%) of tert-butyl 8-[2-(3-[4-[(benzyloxy)carbonyl]piperazin-1-yl]propyl)pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octane-3-carboxylate as a pale yellow oil.
工程7

100mLの丸底フラスコに、メタノール(20mL)中のtert-ブチル8-[2-(3-[4-[(ベンジルオキシ)カルボニル]ピペラジン-1-イル]プロピル)ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレート(2.19g、3.98mmol、1.00当量)を入れた。塩化水素ガスを、室温で1時間撹拌しながら、得られた溶液によって気泡化した。得られた混合物を真空下で濃縮した。これにより、1.79g(92%)のベンジル4-[3-(4-[3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)プロピル]ピペラジン-1-カルボキシレート塩酸塩を淡い黄色の固体として得た。 A 100 mL round bottom flask was charged with tert-butyl 8-[2-(3-[4-[(benzyloxy)carbonyl]piperazin-1-yl]propyl)pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octane-3-carboxylate (2.19 g, 3.98 mmol, 1.00 equiv.) in methanol (20 mL). Hydrogen chloride gas was bubbled through the resulting solution while stirring at room temperature for 1 h. The resulting mixture was concentrated under vacuum. This afforded 1.79 g (92%) of benzyl 4-[3-(4-[3,8-diazabicyclo[3.2.1]octane-8-yl]pyridin-2-yl)propyl]piperazine-1-carboxylate hydrochloride as a pale yellow solid.
工程8

窒素不活性雰囲気下でパージし、維持した20mLのマイクロ波チューブに、DMSO(10mL)中のベンジル4-[3-(4-[3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)プロピル]ピペラジン-1-カルボキシレート塩酸塩(1.79g、3.68mmol、1.00当量)、4-ブロモ-6-クロロピリダジン-3-アミン(2.47g、11.9mmol、3.00当量)、DIEA(5.0mL)を入れた。得られた溶液を、油浴中で、130℃で16時間撹拌した。次いで、反応物を、水/氷(50mL)を添加することによってクエンチした。得られた溶液を、ジクロロメタン(50mL×3)で抽出し、有機層を合わせて、無水硫酸ナトリウムで乾燥させた。残渣を、ジクロロメタン/メタノール(5/1)で溶出するシリカゲルカラム上に適用した。これにより、1.21g(57%)のベンジル4-(3-[4-[3-(3-アミノ-6-クロロピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル]プロピル)ピペラジン-1-カルボキシレートを茶色の固体として得た。以下のスキームに従って、かつ他の実施例について上に記載される手順を使用して、ベンジル4-(3-[4-[3-(3-アミノ-6-クロロピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル]プロピル)ピペラジン-1-カルボキシレートを、6-[2-(メトキシメトキシ)フェニル]-4-(8-[2-[3-(ピペラジン-1-イル)プロピル]ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-イル)ピリダジン-3-アミン(工程10で使用)に変換した。 A 20 mL microwave tube purged and maintained under nitrogen inert atmosphere was charged with benzyl 4-[3-(4-[3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl)propyl]piperazine-1-carboxylate hydrochloride (1.79 g, 3.68 mmol, 1.00 equiv.), 4-bromo-6-chloropyridazin-3-amine (2.47 g, 11.9 mmol, 3.00 equiv.), DIEA (5.0 mL) in DMSO (10 mL). The resulting solution was stirred in an oil bath at 130 °C for 16 h. The reaction was then quenched by adding water/ice (50 mL). The resulting solution was extracted with dichloromethane (50 mL x 3) and the organic layers were combined and dried over anhydrous sodium sulfate. The residue was applied onto a silica gel column eluting with dichloromethane/methanol (5/1). This gave 1.21 g (57%) of benzyl 4-(3-[4-[3-(3-amino-6-chloropyridazin-4-yl)-3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl]propyl)piperazine-1-carboxylate as a brown solid. According to the scheme below and using the procedures described above for other examples, benzyl 4-(3-[4-[3-(3-amino-6-chloropyridazin-4-yl)-3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl]propyl)piperazine-1-carboxylate was converted to 6-[2-(methoxymethoxy)phenyl]-4-(8-[2-[3-(piperazin-1-yl)propyl]pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octan-3-yl)pyridazin-3-amine (used in step 10).
工程9

50mLの丸底フラスコに、N,N-ジメチルホルムアミド(10mL)中の(2S,4R)-4-ヒドロキシ-1-[2-(3-ヒドロキシ-1,2-オキサゾール-5-イル)-3-メチルブタノイル]-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]ピロリジン-2-カルボキサミド(480mg、1.0mmol、1.0当量)の溶液、炭酸カリウム(300mg、2.2mmol、2.2当量)、および1,2-ジブロモエタン(300mg、1.6mmol、1.6当量)を入れた。得られた溶液を、油浴中で、70℃で2時間撹拌した。次いで、反応を、20mLの水の添加により停止させた。得られた溶液を酢酸エチル(3×30mL)で抽出し、有機層を合わせて、無水硫酸ナトリウムで乾燥させ、真空下で濃縮した。残渣を、酢酸エチル/石油エーテル(1:0)で溶出するシリカゲルカラム上に適用した。これにより、235mg(40%)の(2S,4R)-1-[2-[3-(2-ブロモエトキシ)-1,2-オキサゾール-5-イル]-3-メチルブタノイル]-4-ヒドロキシ-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]ピロリジン-2-カルボキサミドを淡い黄色の固体として得た。 A 50 mL round bottom flask was charged with a solution of (2S,4R)-4-hydroxy-1-[2-(3-hydroxy-1,2-oxazol-5-yl)-3-methylbutanoyl]-N-[[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]pyrrolidine-2-carboxamide (480 mg, 1.0 mmol, 1.0 equiv.) in N,N-dimethylformamide (10 mL), potassium carbonate (300 mg, 2.2 mmol, 2.2 equiv.), and 1,2-dibromoethane (300 mg, 1.6 mmol, 1.6 equiv.). The resulting solution was stirred in an oil bath at 70° C. for 2 hours. The reaction was then quenched by the addition of 20 mL of water. The resulting solution was extracted with ethyl acetate (3×30 mL) and the organic layers were combined, dried over anhydrous sodium sulfate, and concentrated under vacuum. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (1:0). This gave 235 mg (40%) of (2S,4R)-1-[2-[3-(2-bromoethoxy)-1,2-oxazol-5-yl]-3-methylbutanoyl]-4-hydroxy-N-[[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]pyrrolidine-2-carboxamide as a pale yellow solid.
工程10
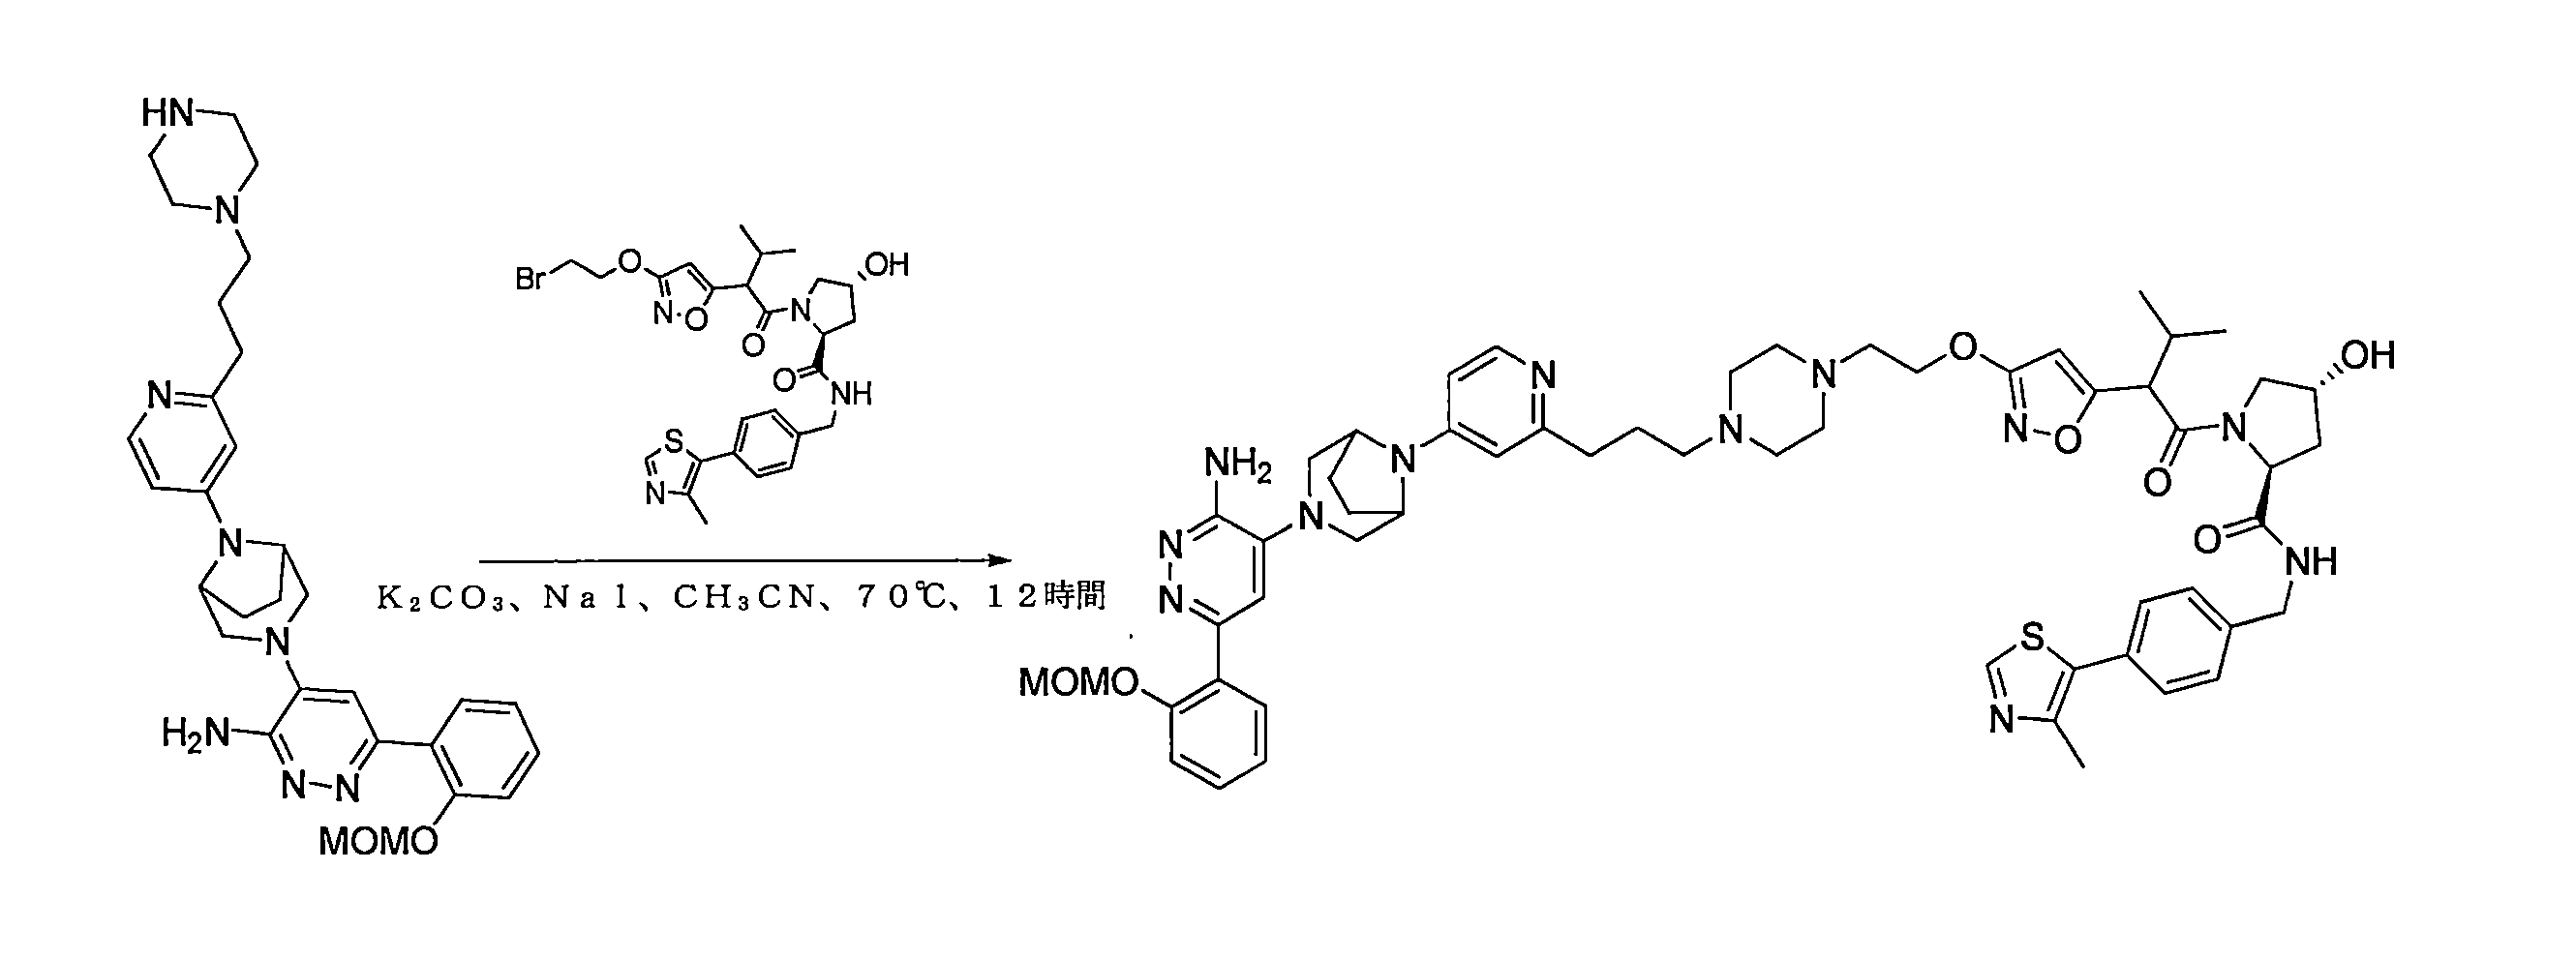
窒素不活性雰囲気下でパージし、維持した20mLのマイクロ波チューブに、アセトニトリル(10mL)中の6-[2-(メトキシメトキシ)フェニル]-4-(8-[2-[3-(ピペラジン-1-イル)プロピル]ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-イル)ピリダジン-3-アミン(92mg、0.17mmol、1.00当量)[以下のスキームに従って、かつ他の実施例について上に記載される手順を使用して、工程8からのベンジル4-(3-[4-[3-(3-アミノ-6-クロロピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル]プロピル)ピペラジン-1-カルボキシレートから調製]、(2S,4R)-1-2-[3-(2-ブロモエトキシ)-1,2-オキサゾール-5-イル]-3-メチルブタノイル-4-ヒドロキシ-N-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチルピロリジン-2-カルボキサミド(100mg、0.17mmol、1.00当量)、炭酸カリウム(70mg、0.51mmol、3.00当量)、NaI(25mg、0.17mmol、1.00当量)を入れた。得られた混合物を12時間、油浴中、70℃で撹拌した。得られた溶液をジクロロメタン(100mL)で抽出し、有機層を合わせた。得られた混合物をブライン(30mL)で洗浄した。混合物を、無水硫酸ナトリウム上で乾燥させ、真空下で濃縮した。これにより、180mg(粗製)の(2S,4R)-1-(2-[3-[2-(4-[3-[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]プロピル]ピペラジン-1-イル)エトキシ]-1,2-オキサゾール-5-イル]-3-メチルブタノイル)-4-ヒドロキシ-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]ピロリジン-2-カルボキサミドを淡い黄色の固体として得た。 In a 20 mL microwave tube purged and maintained under a nitrogen inert atmosphere, 6-[2-(methoxymethoxy)phenyl]-4-(8-[2-[3-(piperazin-1-yl)propyl]pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octan-3-yl)pyridazin-3-amine (92 mg, 0.17 mmol, 1.00 equiv.) [benzyl 4-(3-[4-[3-(3-amino-6-chloropyridazin-4-yl)-3,8-diamine from step 8 according to the scheme below and using the procedures described above for other examples] was added to acetonitrile (10 mL). -diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl]propyl)piperazine-1-carboxylate], (2S,4R)-1-2-[3-(2-bromoethoxy)-1,2-oxazol-5-yl]-3-methylbutanoyl-4-hydroxy-N-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methylpyrrolidine-2-carboxamide (100 mg, 0.17 mmol, 1.00 equiv.), potassium carbonate (70 mg, 0.51 mmol, 3.00 equiv.), and NaI (25 mg, 0.17 mmol, 1.00 equiv.) were added to the reaction mixture. The mixture was stirred at 70° C. in an oil bath for 12 hours. The resulting solution was extracted with dichloromethane (100 mL) and the organic layers were combined. The resulting mixture was washed with brine (30 mL). The mixture was dried over anhydrous sodium sulfate and concentrated under vacuum. This gave 180 mg (crude) of (2S,4R)-1-(2-[3-[2-(4-[3-[4-(3-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl]propyl]piperazin-1-yl)ethoxy]-1,2-oxazol-5-yl]-3-methylbutanoyl)-4-hydroxy-N-[[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]pyrrolidine-2-carboxamide as a pale yellow solid.
以下のスキームに従って、かつ他の実施例について上に記載されるものと類似の手順を使用して、(2S,4R)-1-(2-[3-[2-(4-[3-[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]プロピル]ピペラジン-1-イル)エトキシ]-1,2-オキサゾール-5-イル]-3-メチルブタノイル)-4-ヒドロキシ-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]ピロリジン-2-カルボキサミドを、最終化合物(2S,4R)-1-[(2S)-2-[3-(2-[4-[3-(4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)プロピル]ピペラジン-1-イル]エトキシ)-1,2-オキサゾール-5-イル]-3-メチルブタノイル]-4-ヒドロキシ-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]ピロリジン-2-カルボキサミドおよび(2S,4R)-1-[(2R)-2-[3-(2-[4-[3-(4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)プロピル]ピペラジン-1-イル]エトキシ)-1,2-オキサゾール-5-イル]-3-メチルブタノイル]-4-ヒドロキシ-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]ピロリジン-2-カルボキサミドに変換した。
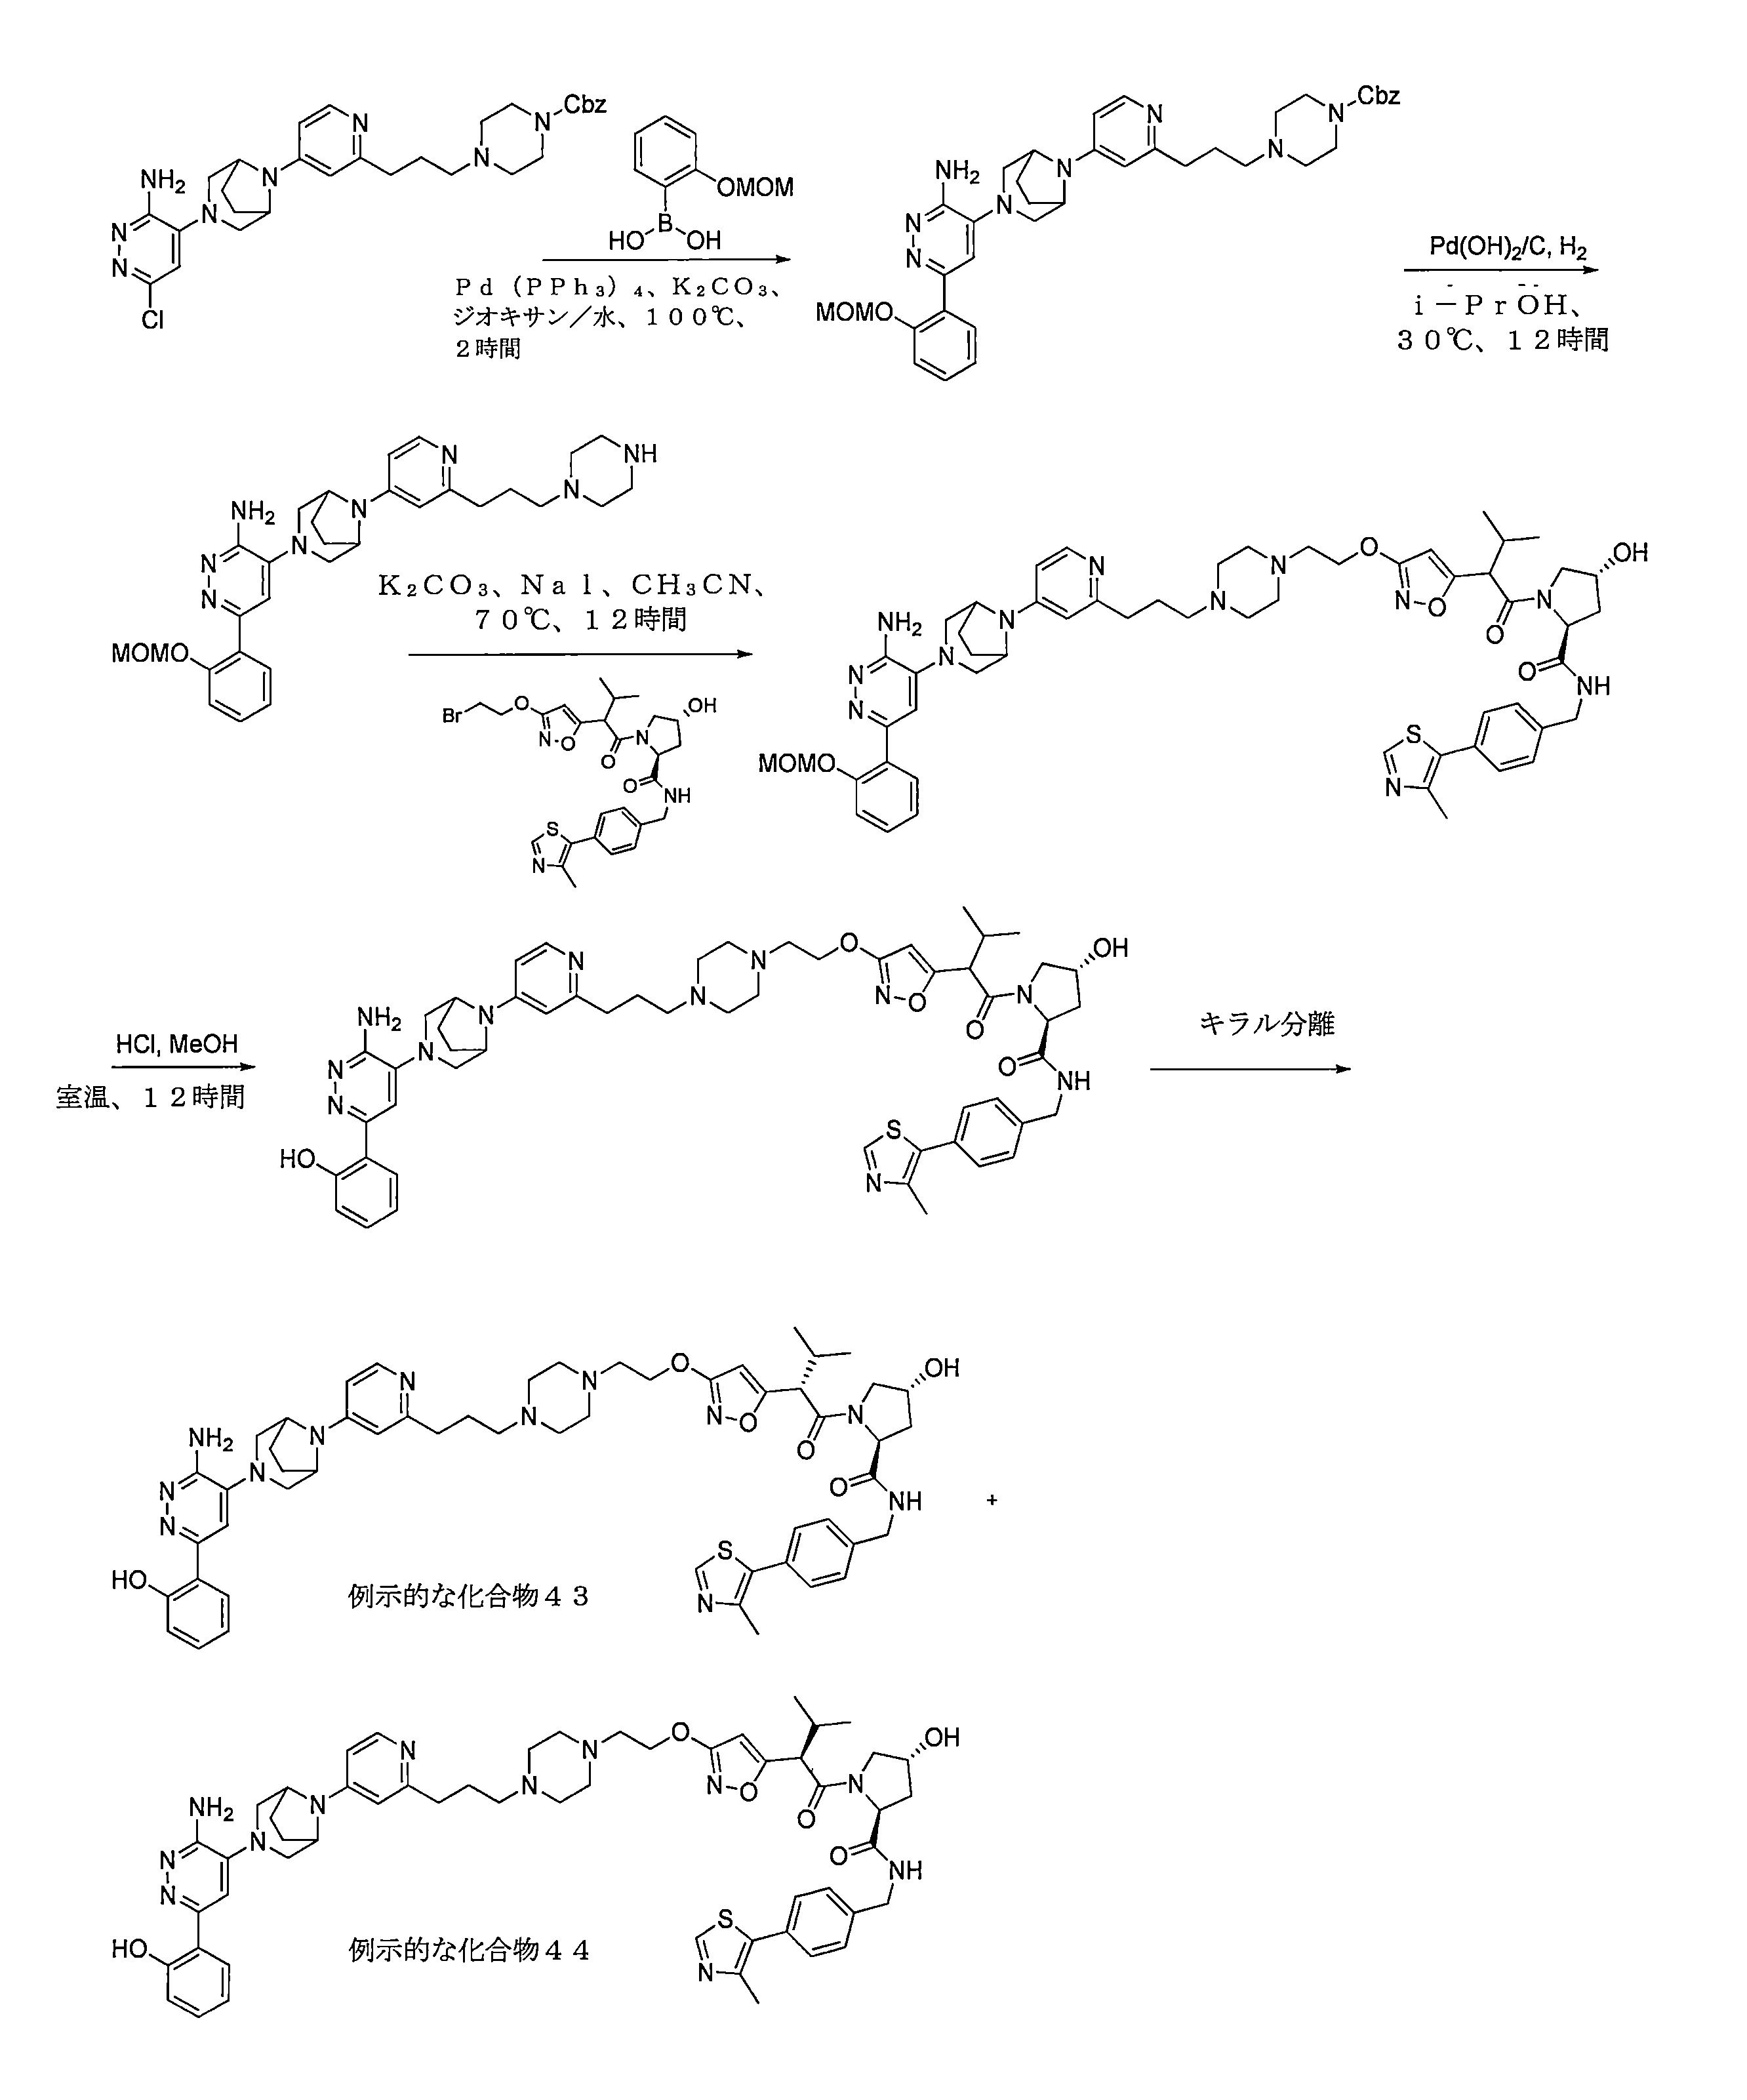
例示的な化合物40の例示的な合成
例示的な化合物40を、上記の他の実施例について記載される手順、ならびに当業者によって理解される手順を使用して、以下のスキームに従って調製した。

例示的な化合物41および例示的な化合物42の例示的な合成
例示的な化合物41および42を、他の例示的な化合物について上に記載される手順を使用して、以下のスキームに従って調製した。

例示的な化合物45の例示的な合成
工程1
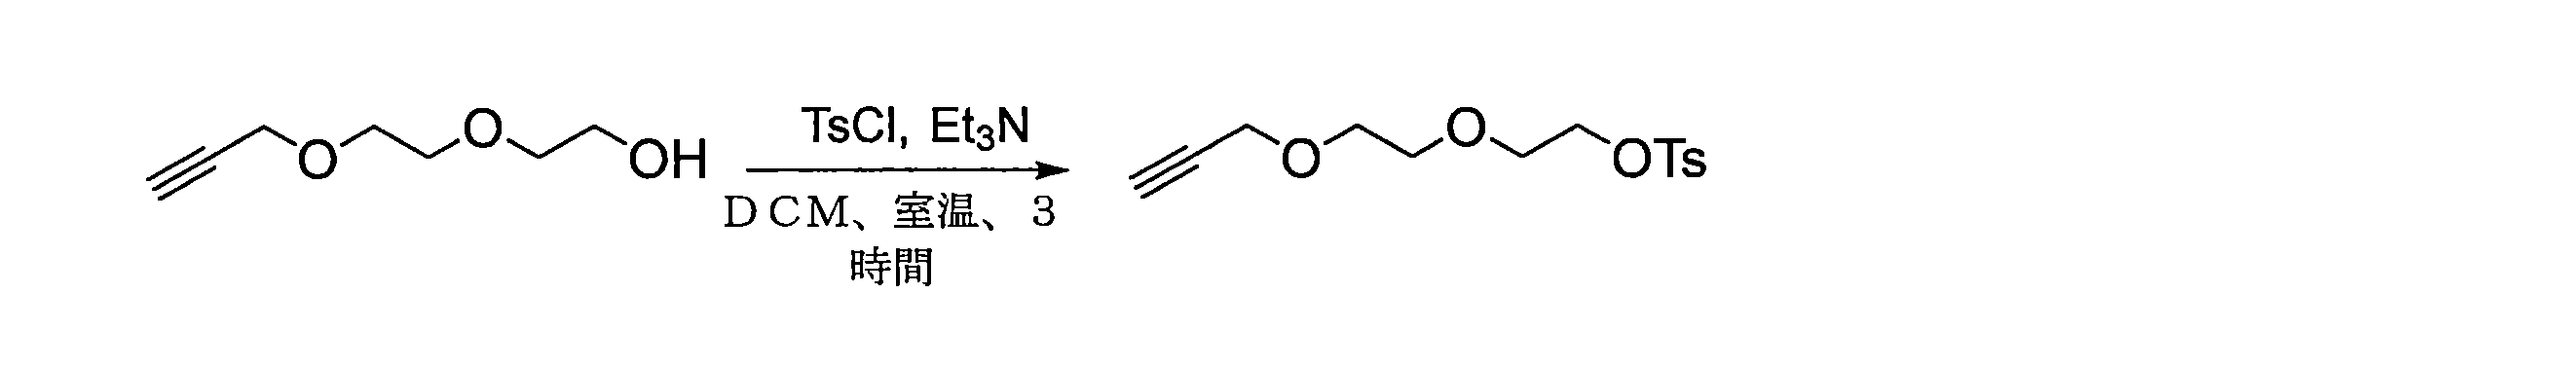
250mLの丸底フラスコに、ジクロロメタン(80mL)中の2-[2-(プロパ-2-イン-1-イルオキシ)エトキシ]エタン-1-オール(1g、6.94mmol、1.00当量)の溶液、トリエチルアミン(2.8g、27.67mmol、3.00当量)、4-ジメチルアミノピリジン(270mg、2.21mmol、0.30当量)、および4-トルエンスルホニルクロリド(1.9g、9.97mmol、1.50当量)を入れた。得られた溶液を室温で12時間撹拌した。得られた混合物を真空下で濃縮した。残渣を、酢酸エチル/石油エーテル(1:2)で溶出するシリカゲルカラム上に適用した。収集した画分をまとめ、真空下で濃縮した。これにより、1.4g(68%)の2-[2-(プロパ-2-イン-1-イルオキシ)エトキシ]エチル4-メチルベンゼン-1-スルホネートを黄色の油として得た。 A 250 mL round-bottom flask was charged with a solution of 2-[2-(prop-2-yn-1-yloxy)ethoxy]ethan-1-ol (1 g, 6.94 mmol, 1.00 equiv.) in dichloromethane (80 mL), triethylamine (2.8 g, 27.67 mmol, 3.00 equiv.), 4-dimethylaminopyridine (270 mg, 2.21 mmol, 0.30 equiv.), and 4-toluenesulfonyl chloride (1.9 g, 9.97 mmol, 1.50 equiv.). The resulting solution was stirred at room temperature for 12 h. The resulting mixture was concentrated under vacuum. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (1:2). The collected fractions were combined and concentrated under vacuum. This yielded 1.4 g (68%) of 2-[2-(prop-2-yn-1-yloxy)ethoxy]ethyl 4-methylbenzene-1-sulfonate as a yellow oil.
工程2

25mLの密封チューブに、N,N-ジメチルホルムアミド(10mL)中の2-[2-(プロパ-2-イン-1-イルオキシ)エトキシ]エチル4-メチルベンゼン-1-スルホネート(1.4g、4.69mmol、1.00当量)の溶液、炭酸カリウム(1.6g、11.58mmol、3.00当量)、ヨウ化ナトリウム(280mg、0.50当量)、およびtert-ブチル2-(ピペラジン-1-イル)酢酸塩(898mg、4.48mmol、1.20当量)を入れた。得られた溶液を、油浴中で、70℃で12時間撹拌した。得られた溶液を酢酸エチルで抽出し、有機層を合わせた。得られた混合物をブラインで洗浄し、真空下で濃縮した。残渣を、酢酸エチル/石油エーテル(1:1)で溶出するシリカゲルカラム上に適用した。収集した画分をまとめ、真空下で濃縮した。これにより、910mg(59%)のtert-ブチル2-(4-[2-[2-(プロパ-2-イン-1-イルオキシ)エトキシ]エチル]ピペラジン-1-イル)酢酸塩を黄色の油として得た。 A 25 mL sealed tube was charged with a solution of 2-[2-(prop-2-yn-1-yloxy)ethoxy]ethyl 4-methylbenzene-1-sulfonate (1.4 g, 4.69 mmol, 1.00 equiv.) in N,N-dimethylformamide (10 mL), potassium carbonate (1.6 g, 11.58 mmol, 3.00 equiv.), sodium iodide (280 mg, 0.50 equiv.), and tert-butyl 2-(piperazin-1-yl)acetate (898 mg, 4.48 mmol, 1.20 equiv.). The resulting solution was stirred in an oil bath at 70° C. for 12 hours. The resulting solution was extracted with ethyl acetate and the organic layers were combined. The resulting mixture was washed with brine and concentrated under vacuum. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (1:1). The collected fractions were combined and concentrated under vacuum. This gave 910 mg (59%) of tert-butyl 2-(4-[2-[2-(prop-2-yn-1-yloxy)ethoxy]ethyl]piperazin-1-yl)acetate as a yellow oil.
工程3

25mLの密封チューブに、テトラヒドロフラン(10mL)中のtert-ブチル2-(4-[2-[2-(プロパ-2-イン-1-イルオキシ)エトキシ]エチル]ピペラジン-1-イル)酢酸塩(910mg、2.79mmol、1.00当量)の溶液、ピナコールボラン(630mg、2.00当量)、ZrCp2HCl(70mg、0.10当量)、およびトリエチルアミン(1mL、3.00当量)を入れた。得られた溶液を、油浴中で、80℃で12時間撹拌した。得られた混合物を真空下で濃縮した。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。収集した画分をまとめ、真空下で濃縮した。これにより、720mg(57%)のtert-ブチル2-[4-[2-(2-[[(2E)-3-(テトラメチル-1,3,2-ジオキサボロラン-2-イル)プロパ-2-エン-1-イル]オキシ]エトキシ)エチル]ピペラジン-1-イル]酢酸塩を黄色の油として得た。 A 25 mL sealed tube was charged with a solution of tert-butyl 2-(4-[2-[2-(prop-2-yn-1-yloxy)ethoxy]ethyl]piperazin-1-yl)acetate (910 mg, 2.79 mmol, 1.00 equiv.) in tetrahydrofuran (10 mL), pinacolborane (630 mg, 2.00 equiv.), ZrCp2HCl (70 mg, 0.10 equiv.), and triethylamine (1 mL, 3.00 equiv.). The resulting solution was stirred in an oil bath at 80°C for 12 h. The resulting mixture was concentrated under vacuum. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (10:1). The collected fractions were combined and concentrated under vacuum. This gave 720 mg (57%) of tert-butyl 2-[4-[2-(2-[[(2E)-3-(tetramethyl-1,3,2-dioxaborolan-2-yl)prop-2-en-1-yl]oxy]ethoxy)ethyl]piperazin-1-yl]acetate as a yellow oil.
工程4

100mLの丸底フラスコに、ジメチルスルホキシド(40mL)中のtert-ブチル3,8-ジアザビシクロ[3.2.1]オクタン-8-カルボキシレート(2.0g、9.42mmol、1.00当量)の溶液、N,N-ジイソプロピルエチルアミン(714.0mg、5.52mmol、4.00当量)、4-ブロモ-6-クロロピリダジン-3-アミン(3.0g、14.39mmol、1.50当量)を入れた。得られた溶液を、油浴中で、130℃で3時間撹拌した。次いで、反応物を、水(90mL)を添加することによってクエンチした。得られた溶液を酢酸エチル(80mL×3)で抽出し、有機層を合わせた。得られた混合物をブライン(100mL×1)で洗浄した。混合物を、無水硫酸ナトリウム上で乾燥させ、真空下で濃縮した。残渣を、ジクロロメタン/メタノール(10/1)とともにシリカゲルカラムに充填させた。収集した画分をまとめ、真空下で濃縮した。これにより、1.7g(53%)のtert-ブチル3-(3-アミノ-6-クロロピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-カルボキシレートを黄色の固体として得た。 A 100 mL round bottom flask was charged with a solution of tert-butyl 3,8-diazabicyclo[3.2.1]octane-8-carboxylate (2.0 g, 9.42 mmol, 1.00 equiv.) in dimethyl sulfoxide (40 mL), N,N-diisopropylethylamine (714.0 mg, 5.52 mmol, 4.00 equiv.), 4-bromo-6-chloropyridazin-3-amine (3.0 g, 14.39 mmol, 1.50 equiv.). The resulting solution was stirred in an oil bath at 130 °C for 3 h. The reaction was then quenched by adding water (90 mL). The resulting solution was extracted with ethyl acetate (80 mL x 3) and the organic layers were combined. The resulting mixture was washed with brine (100 mL x 1). The mixture was dried over anhydrous sodium sulfate and concentrated under vacuum. The residue was loaded onto a silica gel column with dichloromethane/methanol (10/1). The collected fractions were combined and concentrated under vacuum. This gave 1.7 g (53%) of tert-butyl 3-(3-amino-6-chloropyridazin-4-yl)-3,8-diazabicyclo[3.2.1]octane-8-carboxylate as a yellow solid.
工程5

窒素不活性雰囲気下でパージし、維持した250mLの三口丸底フラスコに、ジオキサン/水(50/10mL)中のtert-ブチル3-(3-アミノ-6-クロロピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-カルボキシレート(1.7g、5.00mmol、1.00当量)の溶液、炭酸カリウム(2.1g、15.19mmol、3.00当量)、(2-ヒドロキシフェニル)ボロン酸(1.04g、7.54mmol、1.50当量)、テトラキス(トリフェニルホスフィン)パラジウム(0)(578mg、0.50mmol、0.10当量)を入れた。得られた溶液を2時間、油浴中、100℃で撹拌した。次いで、反応を水(60mL)の添加によりクエンチさせた。得られた溶液を酢酸エチル(40mL×3)で抽出し、有機層を合わせた。得られた混合物をブライン(60mL×1)で洗浄した。混合物を、無水硫酸ナトリウム上で乾燥させ、真空下で濃縮した。残渣を、ジクロロメタン/メタノール(10/1)で溶出するシリカゲルカラム上に適用した。収集した画分をまとめ、真空下で濃縮した。これにより、1.62g(81%)のtert-ブチル3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-カルボキシレートを黄色の固体として得た。 A 250 mL three-neck round bottom flask purged and maintained under nitrogen inert atmosphere was charged with a solution of tert-butyl 3-(3-amino-6-chloropyridazin-4-yl)-3,8-diazabicyclo[3.2.1]octane-8-carboxylate (1.7 g, 5.00 mmol, 1.00 equiv.) in dioxane/water (50/10 mL), potassium carbonate (2.1 g, 15.19 mmol, 3.00 equiv.), (2-hydroxyphenyl)boronic acid (1.04 g, 7.54 mmol, 1.50 equiv.), tetrakis(triphenylphosphine)palladium(0) (578 mg, 0.50 mmol, 0.10 equiv.). The resulting solution was stirred at 100°C in an oil bath for 2 hours. The reaction was then quenched by the addition of water (60 mL). The resulting solution was extracted with ethyl acetate (40 mL x 3) and the organic layers were combined. The resulting mixture was washed with brine (60 mL x 1). The mixture was dried over anhydrous sodium sulfate and concentrated under vacuum. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (10/1). The collected fractions were combined and concentrated under vacuum. This gave 1.62 g (81%) of tert-butyl 3-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octane-8-carboxylate as a yellow solid.
工程6

250mLの丸底フラスコに、ジオキサン(50mL)中のtert-ブチル3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-カルボキシレート(1.62g、4.08mmol、1.00当量)の溶液を入れた。上記に塩化水素(ガス)を導入した。得られた溶液を室温で5時間撹拌した。得られた混合物を真空下で濃縮した。これにより、1.2g(88%)の2-(6-アミノ-5-[3,8-ジアザビシクロ[3.2.1]オクタン-3-イル]ピリダジン-3-イル)フェノール塩酸塩を黄色の固体として得た。 A 250 mL round bottom flask was charged with a solution of tert-butyl 3-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octane-8-carboxylate (1.62 g, 4.08 mmol, 1.00 equiv.) in dioxane (50 mL). Hydrogen chloride (gas) was introduced to the above. The resulting solution was stirred at room temperature for 5 hours. The resulting mixture was concentrated under vacuum. This afforded 1.2 g (88%) of 2-(6-amino-5-[3,8-diazabicyclo[3.2.1]octan-3-yl]pyridazin-3-yl)phenol hydrochloride as a yellow solid.
工程7
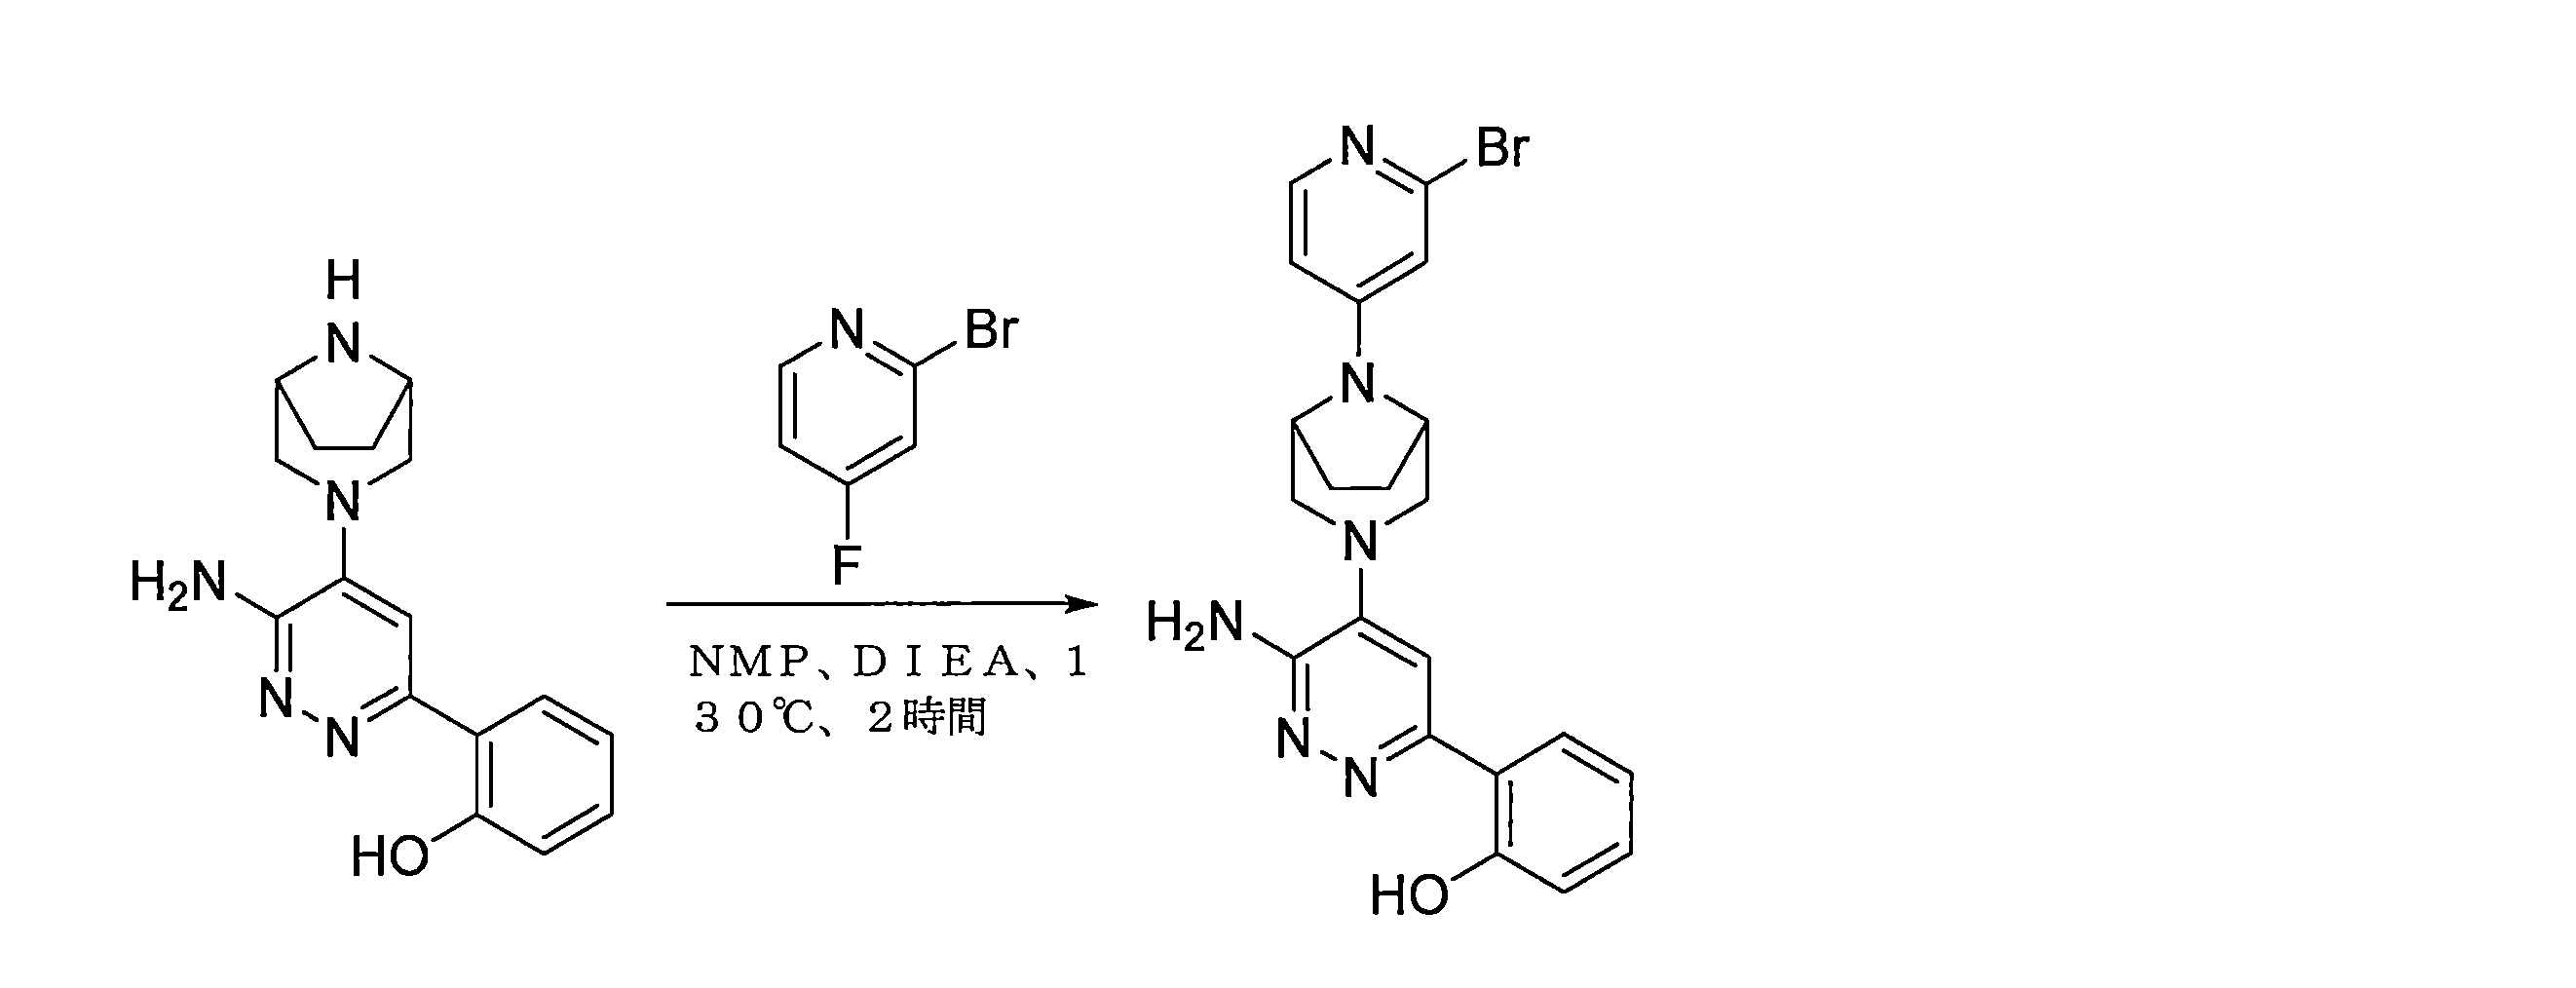
100mLの丸底フラスコに、1-メチル-2-ピロリジノン(30mL)中の2-(6-アミノ-5-[3,8-ジアザビシクロ[3.2.1]オクタン-3-イル]ピリダジン-3-イル)フェノール(400mg、1.35mmol、1.00当量)の溶液、N,N-ジイソプロピルエチルアミン(693mg、5.36mmol、4.00当量)、2-ブロモ-4-フルオロピリジン(355.0mg、2.02mmol、1.50当量)を入れた。得られた溶液を、油浴中で、130℃で2時間撹拌した。次いで、反応物を、水(40mL)を添加することによってクエンチした。得られた溶液を酢酸エチル(30mL×3)で抽出し、有機層を合わせた。得られた混合物をブライン(40mL×1)で洗浄した。混合物を、無水硫酸ナトリウム上で乾燥させ、真空下で濃縮した。残渣を、酢酸エチル/石油エーテル(4/1)で溶出するシリカゲルカラム上に適用した。収集した画分をまとめ、真空下で濃縮した。これにより、410mg(67%)の2-[6-アミノ-5-[8-(2-ブロモピリジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-3-イル]ピリダジン-3-イル]フェノールを黄色の固体として得た。 A 100 mL round bottom flask was charged with a solution of 2-(6-amino-5-[3,8-diazabicyclo[3.2.1]octan-3-yl]pyridazin-3-yl)phenol (400 mg, 1.35 mmol, 1.00 equiv.) in 1-methyl-2-pyrrolidinone (30 mL), N,N-diisopropylethylamine (693 mg, 5.36 mmol, 4.00 equiv.), 2-bromo-4-fluoropyridine (355.0 mg, 2.02 mmol, 1.50 equiv.). The resulting solution was stirred in an oil bath at 130° C. for 2 hours. The reaction was then quenched by adding water (40 mL). The resulting solution was extracted with ethyl acetate (30 mL×3) and the organic layers were combined. The resulting mixture was washed with brine (40 mL×1). The mixture was dried over anhydrous sodium sulfate and concentrated under vacuum. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (4/1). The collected fractions were combined and concentrated under vacuum. This gave 410 mg (67%) of 2-[6-amino-5-[8-(2-bromopyridin-4-yl)-3,8-diazabicyclo[3.2.1]octan-3-yl]pyridazin-3-yl]phenol as a yellow solid.
工程8

20mLの密封チューブに、ジオキサン/水(5:1mL)中のtert-ブチル2-[4-[2-(2-[[(2E)-3-(テトラメチル-1,3,2-ジオキサボロラン-2-イル)プロパ-2-エン-1-イル]オキシ]エトキシ)エチル]ピペラジン-1-イル]酢酸塩(628mg、1.38mmol、1.00当量)の溶液、テトラキス(トリフェニルホスフィンホスフィン)パラジウム(108mg、0.15mmol、0.10当量)、炭酸カリウム(286mg、2.07mmol、1.50当量)、2-(6-アミノ-5-((1R,5S)-8-(2-ブロモピリジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-3-イル)ピリダジン-3-イル)フェノール(500mg、1.10mmol、0.80当量)を入れた。得られた溶液を、油浴中で、80℃で2時間撹拌した。得られた混合物を真空下で濃縮した。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。収集した画分をまとめ、真空下で濃縮した。これにより、540mg(56%)のtert-ブチル2-[4-[2-(2-[[(2E)-3-(4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)プロパ-2-エン-1-イル]オキシ]エトキシ)エチル]ピペラジン-1-イル]酢酸塩を黄色の固体として得た。 In a 20 mL sealed tube, add a solution of tert-butyl 2-[4-[2-(2-[[(2E)-3-(tetramethyl-1,3,2-dioxaborolan-2-yl)prop-2-en-1-yl]oxy]ethoxy)ethyl]piperazin-1-yl]acetate (628 mg, 1.38 mmol, 1.00 equiv.) in dioxane/water (5:1 mL), tetrakis(triphenylphosphinephosphine) (1R,5S)-8-(2-bromopyridin-4-yl)-3,8-diazabicyclo[3.2.1]octan-3-yl)pyridazin-3-yl)phenol (500 mg, 1.10 mmol, 0.80 equiv.) was added to a 500 mL flask. The resulting solution was stirred in an oil bath at 80° C. for 2 hours. The resulting mixture was concentrated under vacuum. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (10:1). The collected fractions were combined and concentrated under vacuum. This yielded 540 mg (56%) of tert-butyl 2-[4-[2-(2-[[(2E)-3-(4-[3-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl)prop-2-en-1-yl]oxy]ethoxy)ethyl]piperazin-1-yl]acetate as a yellow solid.
工程9
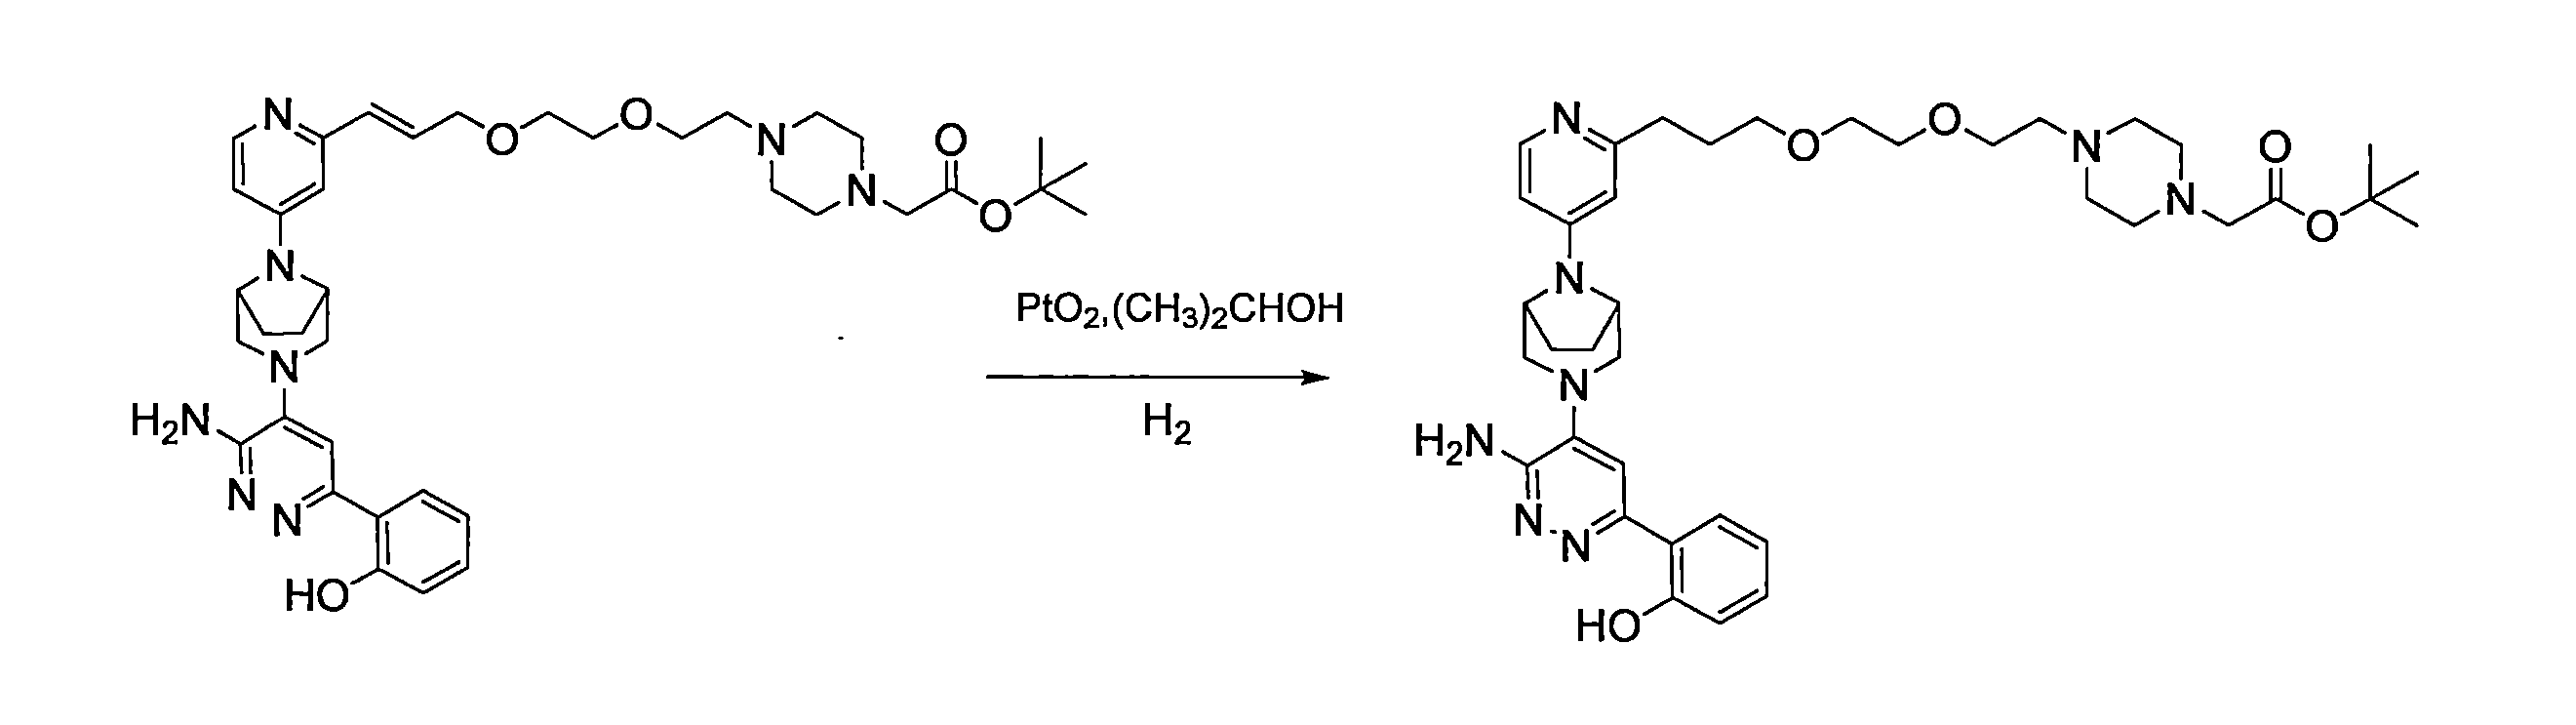
100mLの丸底フラスコに、2-プロパノール(30mL)中のtert-ブチル2-[4-[2-(2-[[(2E)-3-(4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)プロパ-2-エン-1-イル]オキシ]エトキシ)エチル]ピペラジン-1-イル]酢酸塩(200mg、0.29mmol、1.00当量)の溶液およびPtO2(10mg、0.10当量)を入れた。得られた溶液を、水素雰囲気下で、室温で12時間撹拌した。得られた溶液を50mLの2-プロパノールで希釈し、次いで、真空下で濃縮した。これにより、190mg(95%)のtert-ブチル2-[4-(2-[2-[3-(4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)プロポキシ]エトキシ]エチル)ピペラジン-1-イル]酢酸塩を黄色の固体として得た。 A 100 mL round bottom flask was charged with a solution of tert-butyl 2-[4-[2-(2-[[(2E)-3-(4-[3-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl)prop-2-en-1-yl]oxy]ethoxy)ethyl]piperazin-1-yl]acetate (200 mg, 0.29 mmol, 1.00 equiv.) in 2-propanol (30 mL) and PtO2 (10 mg, 0.10 equiv.). The resulting solution was stirred at room temperature under a hydrogen atmosphere for 12 h. The resulting solution was diluted with 50 mL of 2-propanol and then concentrated under vacuum. This yielded 190 mg (95%) of tert-butyl 2-[4-(2-[2-[3-(4-[3-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl)propoxy]ethoxy]ethyl)piperazin-1-yl]acetate as a yellow solid.
工程10
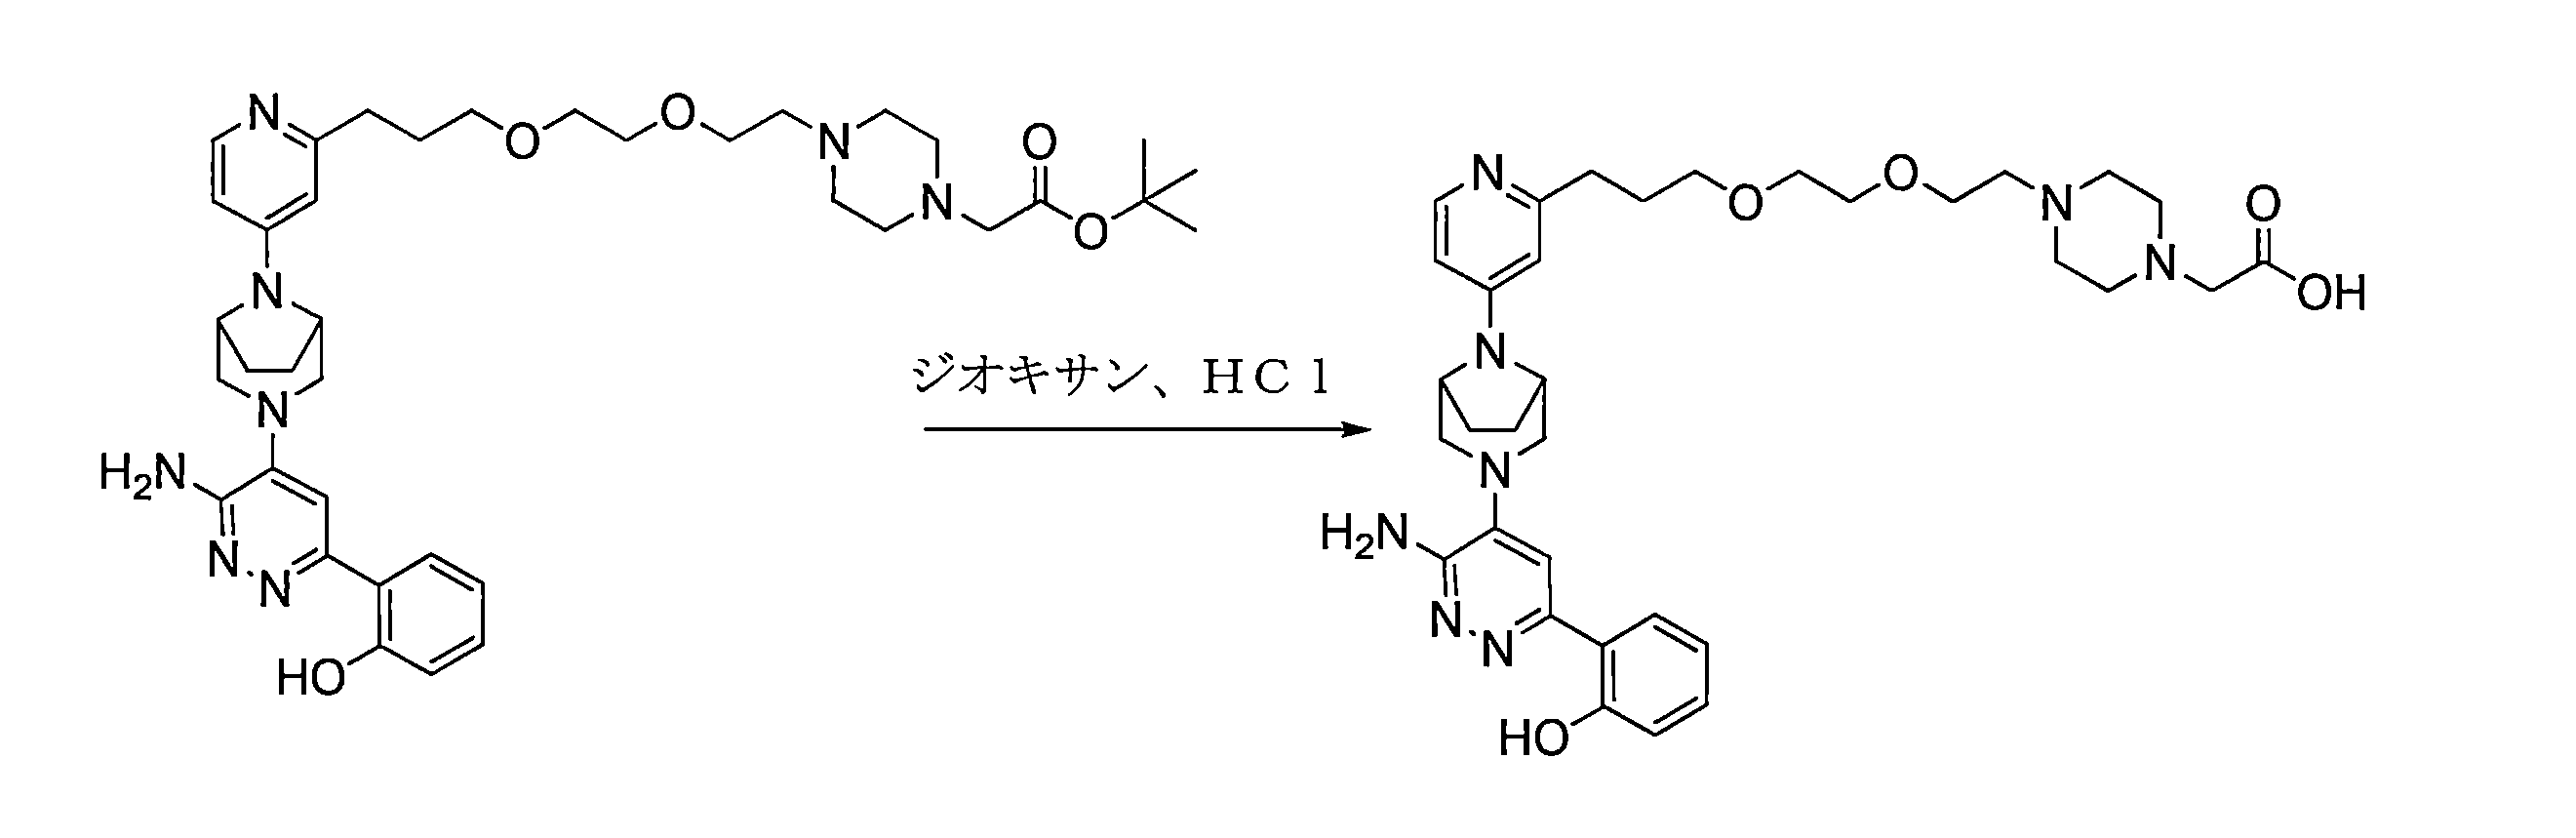
100mLの丸底フラスコに、ジオキサン(20mL)中のtert-ブチル2-[4-(2-[2-[3-(4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)プロポキシ]エトキシ]エチル)ピペラジン-1-イル]酢酸塩(190mg、0.27mmol、1.00当量)の溶液を入れた。上記に塩化水素(ガス)を導入した。得られた溶液を室温で2時間撹拌した。得られた混合物を真空下で濃縮した。これにより、160mg(92%)の2-[4-(2-[2-[3-(4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)プロポキシ]エトキシ]エチル)ピペラジン-1-イル]酢酸を黄色の固体として得た。 A 100 mL round bottom flask was charged with a solution of tert-butyl 2-[4-(2-[2-[3-(4-[3-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl)propoxy]ethoxy]ethyl)piperazin-1-yl]acetate (190 mg, 0.27 mmol, 1.00 equiv.) in dioxane (20 mL). Hydrogen chloride (gas) was introduced to the above. The resulting solution was stirred at room temperature for 2 hours. The resulting mixture was concentrated under vacuum. This gave 160 mg (92%) of 2-[4-(2-[2-[3-(4-[3-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl)propoxy]ethoxy]ethyl)piperazin-1-yl]acetic acid as a yellow solid.
工程11
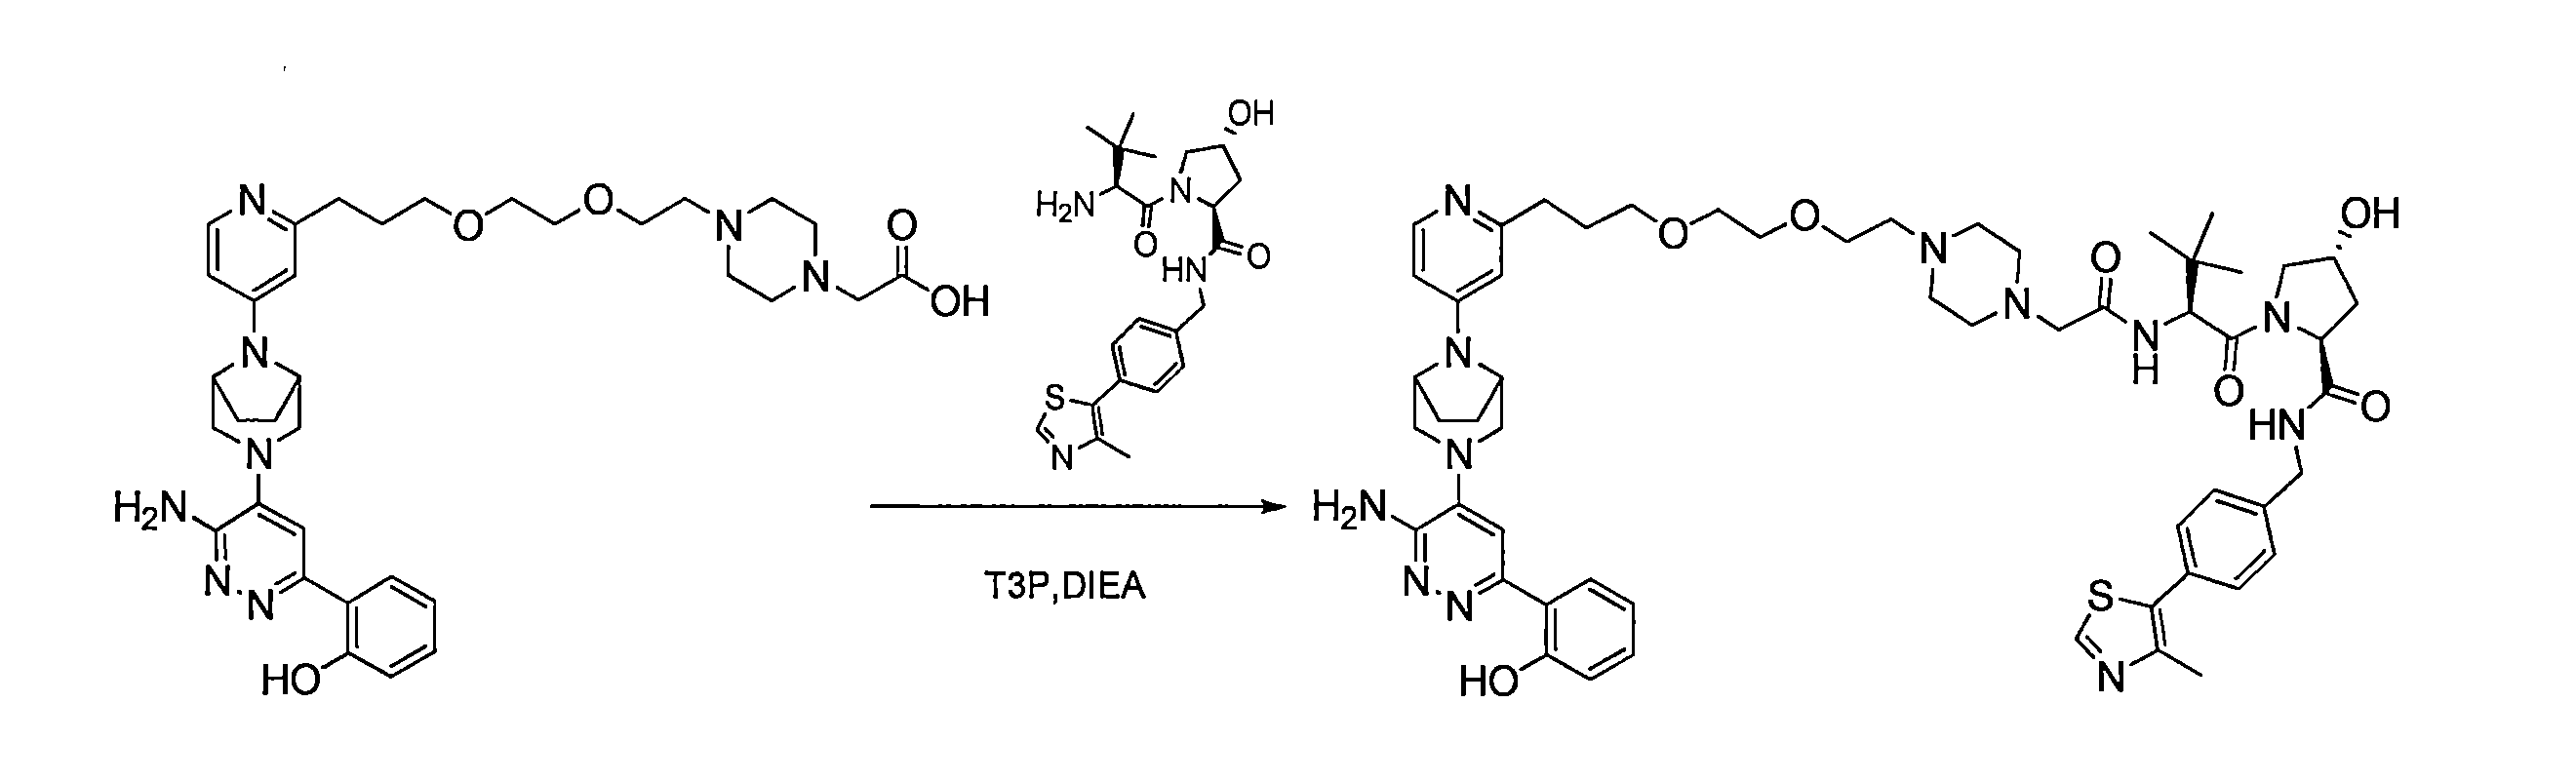
100mLの丸底フラスコに、2-[4-(2-2-[3-(4-3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イルピリジン-2-イル)プロポキシ]エトキシエチル)ピペラジン-1-イル]酢酸(160mg、0.25mmol、1.00当量)、N,N-ジイソプロピルエチルアミン(0.3mL、0.30当量)、アセトニトリル(20mL)中の(2S,4R)-1-[(2S)-2-アミノ-3,3-ジメチルブタノイル]-4-ヒドロキシ-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]ピロリジン-2-カルボキサミド塩酸塩(139mg、0.30mmol、1.20当量)の溶液、およびT3P(188mg、3.00当量)を入れた。得られた溶液を室温で2時間撹拌した。得られた混合物を真空下で濃縮した。粗生成物(100mL)を、以下の条件を用いて、分取HPLCによって精製した。カラム、XBridge Prep C18 OBDカラム、5um、19*150mm;移動相、水(10mmol/Lの重炭酸アンモニウム)およびアセトニトリル(26.0%アセトニトリル、8分間で最大49.0%);検出器、UV254nm。これにより、28.5mg(11%)の(2S,4R)-1-[(2S)-2-[2-[4-(2-[2-[3-(4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)プロポキシ]エトキシ]エチル)ピペラジン-1-イル]アセトアミド]-3,3-ジメチルブタノイル]-4-ヒドロキシ-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]ピロリジン-2-カルボキサミドを白色の固体として得た。 In a 100 mL round-bottom flask, add 2-[4-(2-2-[3-(4-3-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-ylpyridin-2-yl)propoxy]ethoxyethyl)piperazin-1-yl]acetic acid (160 mg, 0.25 mmol, 1.00 equiv.), N,N-diisopropylethylamine (0.3 m A solution of (2S,4R)-1-[(2S)-2-amino-3,3-dimethylbutanoyl]-4-hydroxy-N-[[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]pyrrolidine-2-carboxamide hydrochloride (139 mg, 0.30 mmol, 1.20 equiv.) in acetonitrile (20 mL), and T3P (188 mg, 3.00 equiv.) was charged. The resulting solution was stirred at room temperature for 2 h. The resulting mixture was concentrated under vacuum. The crude product (100 mL) was purified by preparative HPLC using the following conditions: Column, XBridge Prep C18 OBD column, 5um, 19*150mm; mobile phase, water (10mmol/L ammonium bicarbonate) and acetonitrile (26.0% acetonitrile, max 49.0% in 8min); detector, UV 254nm. This gave 28.5 mg (11%) of (2S,4R)-1-[(2S)-2-[2-[4-(2-[2-[3-(4-[3-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl)propoxy]ethoxy]ethyl)piperazin-1-yl]acetamide]-3,3-dimethylbutanoyl]-4-hydroxy-N-[[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]pyrrolidine-2-carboxamide as a white solid.
1H NMR(300MHz、CD3OD):δ8.88~8.66(m、1H)、8.09~7.88(m、1H)、7.76~7.57(m、1H)、7.52~7.01(m、6H)、6.92~6.62(m、4H)、4.66~4.21(m、6H)、3.94~3.72(m、2H)、3.69~3.33(m、12H)、3.14~2.89(m、4H)、2.81~2.34(m、14H)、2.32~1.78(m、8H)、1.32~0.74(m、12H)。 1H NMR (300MHz, CD3OD): δ 8.88-8.66 (m, 1H), 8.09-7.88 (m, 1H), 7.76-7.57 (m, 1H), 7.52-7.01 (m, 6H), 6.92-6.62 (m, 4H), 4.66-4.21 (m, 6H), 3.94-3.72 (m, 2H), 3.69-3.33 (m, 12H), 3.14-2.89 (m, 4H), 2.81-2.34 (m, 14H), 2.32-1.78 (m, 8H), 1.32-0.74 (m, 12H).
例示的な化合物46および例示的な化合物47の例示的な合成
工程1
50mLの丸底フラスコに、N,N-ジメチルホルムアミド(20mL)中の2-(3-メトキシ-1,2-オキサゾール-5-イル)-3-メチルブタン酸(2.0g、10.0mmol、1.0当量)の溶液、N,N,N’,N’-テトラメチル-O-(7-アザベンゾトリアゾール-1-イル)ウロニウムヘキサフルオロホスフェート(4.1g、10.8mmol、1.1当量)、N,N-ジイソプロピルエチルアミン(6.0g、46.4mmol、4.6当量)、および(2S,4R)-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド塩酸塩(3.5g、9.5mmol、1.0当量)を入れた。得られた溶液を室温で30分間撹拌した。次いで、反応を30mLの水の添加によりクエンチさせた。得られた溶液を酢酸エチル(30mL×3)で抽出し、有機層を合わせて、真空下で濃縮した。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。これにより、2.1g(41%)の(2S,4R)-4-ヒドロキシ-1-[2-(3-メトキシ-1,2-オキサゾール-5-イル)-3-メチルブタノイル]-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミドを淡い黄色の固体として得た。 A 50 mL round-bottom flask was charged with a solution of 2-(3-methoxy-1,2-oxazol-5-yl)-3-methylbutanoic acid (2.0 g, 10.0 mmol, 1.0 equiv.) in N,N-dimethylformamide (20 mL), N,N,N',N'-tetramethyl-O-(7-azabenzotriazol-1-yl)uronium hexafluorophosphate (4.1 g, 10.8 mmol, 1.1 equiv.), N,N-diisopropylethylamine (6.0 g, 46.4 mmol, 4.6 equiv.), and (2S,4R)-4-hydroxy-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide hydrochloride (3.5 g, 9.5 mmol, 1.0 equiv.). The resulting solution was stirred at room temperature for 30 min. The reaction was then quenched by the addition of 30 mL of water. The resulting solution was extracted with ethyl acetate (30 mL x 3) and the organic layers were combined and concentrated under vacuum. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (10:1). This gave 2.1 g (41%) of (2S,4R)-4-hydroxy-1-[2-(3-methoxy-1,2-oxazol-5-yl)-3-methylbutanoyl]-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide as a pale yellow solid.
工程2

50mLの丸底フラスコに、(2S,4R)-4-ヒドロキシ-1-[2-(3-メトキシ-1,2-オキサゾール-5-イル)-3-メチルブタノイル]-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(2.1g、4.1mmol、1.0当量)および48%臭化水素水溶液(15mL)を入れた。得られた溶液を、油浴中で、60℃で一晩撹拌した。得られた混合物を、真空下で濃縮して、1.9gの粗生成物を生成した。 A 50 mL round bottom flask was charged with (2S,4R)-4-hydroxy-1-[2-(3-methoxy-1,2-oxazol-5-yl)-3-methylbutanoyl]-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (2.1 g, 4.1 mmol, 1.0 equiv.) and 48% aqueous hydrogen bromide (15 mL). The resulting solution was stirred overnight at 60° C. in an oil bath. The resulting mixture was concentrated under vacuum to yield 1.9 g of crude product.
工程3

50mLの丸底フラスコに、CH3CN(20mL)中の(2S,4R)-4-ヒドロキシ-1-[2-(3-ヒドロキシ-1,2-オキサゾール-5-イル)-3-メチルブタノイル]-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(1.9g、3.8mmol、1.0当量)の溶液、炭酸カリウム(3.5g、25.3mmol、6.6当量)、ヨウ化ナトリウム(800.0mg、5.3mmol、1.4当量)、および1,2-ジブロモエタン(1g、5.3mmol、1.4当量)を入れた。得られた溶液を、油浴中で、60℃で一晩撹拌した。得られた混合物を真空下で濃縮した。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。これにより、1.1g(48%)の(2S,4R)-1-[2-[3-(2-ブロモエトキシ)-1,2-オキサゾール-5-イル]-3-メチルブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミドを白色の固体として得た。 A 50 mL round-bottom flask was charged with a solution of (2S,4R)-4-hydroxy-1-[2-(3-hydroxy-1,2-oxazol-5-yl)-3-methylbutanoyl]-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (1.9 g, 3.8 mmol, 1.0 equiv.) in CH3CN (20 mL), potassium carbonate (3.5 g, 25.3 mmol, 6.6 equiv.), sodium iodide (800.0 mg, 5.3 mmol, 1.4 equiv.), and 1,2-dibromoethane (1 g, 5.3 mmol, 1.4 equiv.). The resulting solution was stirred overnight at 60 °C in an oil bath. The resulting mixture was concentrated under vacuum. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (10:1). This gave 1.1 g (48%) of (2S,4R)-1-[2-[3-(2-bromoethoxy)-1,2-oxazol-5-yl]-3-methylbutanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide as a white solid.
工程4

50mLの丸底フラスコに、CH3CN(20mL)中の(2S,4R)-1-[2-[3-(2-ブロモエトキシ)-1,2-オキサゾール-5-イル]-3-メチルブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(200mg、0.3mmol、1.0当量)の溶液、炭酸カリウム(400mg、2.9mmol、8.8当量)、ヨウ化ナトリウム(80mg、0.53mmol、1.6当量)、および6-[2-(メトキシメトキシ)フェニル]-4-(8-2-[2-(ピペラジン-1-イル)エトキシ]ピリジン-4-イル-3,8-ジアザビシクロ[3.2.1]オクタン-3-イル)ピリダジン-3-アミン(190mg、0.4mmol、1.1当量)[他の実施例について上に記載されるように調製]を入れた。得られた溶液を、油浴中で、60℃で一晩撹拌した。得られた混合物を真空下で濃縮した。残渣を、ジクロロメタン/メタノール(5:1)で溶出するシリカゲルカラム上に適用した。これにより、180mg(51%)の(2S,4R)-1-[2-(3-[2-[4-(2-[[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]オキシ]エチル)ピペラジン-1-イル]エトキシ]-1,2-オキサゾール-5-イル)-3-メチルブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミドを淡い黄色の固体として得た。 In a 50 mL round bottom flask, add a solution of (2S,4R)-1-[2-[3-(2-bromoethoxy)-1,2-oxazol-5-yl]-3-methylbutanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (200 mg, 0.3 mmol, 1.0 equiv.) in CH3CN (20 mL), potassium carbonate (400 mg, 2.9 mmol, 8.8 equiv.), sodium iodide (80 mg, 0.53 mmol, 1.6 equiv.), and 6-[2-(methoxymethoxy)phenyl]-4-(8-2-[2-(piperazin-1-yl)ethoxy]pyridin-4-yl-3,8-diazabicyclo[3.2.1]octan-3-yl)pyridazin-3-amine (190 mg, 0.4 mmol, 1.1 equiv.) [prepared as described above for other examples]. The resulting solution was stirred overnight at 60° C. in an oil bath. The resulting mixture was concentrated under vacuum. The residue was applied onto a silica gel column eluting with dichloromethane/methanol (5:1). This gave 180 mg (51%) of (2S,4R)-1-[2-(3-[2-[4-(2-[[4-(3-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl]oxy]ethyl)piperazin-1-yl]ethoxy]-1,2-oxazol-5-yl)-3-methylbutanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide as a pale yellow solid.
工程5
50mLの丸底フラスコに、メタノール(5mL)中の(2S,4R)-1-[2-(3-[2-[4-(2-[[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]オキシ]エチル)ピペラジン-1-イル]エトキシ]-1,2-オキサゾール-5-イル)-3-メチルブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(180mg、0.17mmol、1.0当量)の溶液、および塩化水素(0.068mLのジオキサン中の4M溶液、0.27mmol、1.6当量)を入れた。得られた溶液を室温で2時間撹拌した。得られた混合物を真空下で濃縮した。粗生成物を、以下の条件を用いて、キラル分取HPLCによって精製した。カラム;移動相、カラム:CHIRALPAK ID-03、2.0cm I.D*25cm L(5um);移動相A:MTBE(0.2%IPA)--HPLC、移動相B:メタノール--HPLC、流速:18mL/分、勾配:18分間で50B~50B、254/220nm。これにより、46.7mgの(2S,4R)-1-[(2S)-2-[3-[2-(4-[2-[(4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)オキシ]エチル]ピペラジン-1-イル)エトキシ]-1,2-オキサゾール-5-イル]-3-メチルブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミドを白色の固体[1H NMR(300MHz、DMSO-d6)δ14.10(s、1H)、8.99~8.83(m、1H)、8.25(d、J=7.8Hz、1H)、7.92~7.83(m、2H)、7.75~7.12(m、6H)、6.81(td、J=8.4、2.2Hz、2H)、6.49(dd、J=6.1、2.0Hz、1H)、6.15~5.93(m、4H)、5.06(d、J=3.6Hz、1H)、4.98~4.79(m、1H)、4.38(t、J=7.7Hz、3H)、4.19(dt、J=21.0、5.6Hz、5H)、3.98(m、1H)、3.71(d、J=8.6Hz、2H)、3.57~3.34(m、2H)、3.33~3.15(m、2H)、2.97(d、J=11.6Hz、4H)、2.57~2.40(d、J=8.0Hz、11H)、2.31~2.07(m、1H)、2.06~1.97(m、2H)、1.91~1.66(m、3H)、1.42(d、J=7.0Hz、1H)、1.31(d、J=7.0Hz、3H)、0.92~0.67(m、6H)]として、および54.4mgの(2S,4R)-1-[(2R)-2-[3-[2-(4-[2-[(4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)オキシ]エチル]ピペラジン-1-イル)エトキシ]-1,2-オキサゾール-5-イル]-3-メチルブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミドを白色の固体[1H NMR(300MHz、DMSO-d6)δ14.11(s、1H)、8.95(s、1H)、8.37(d、J=7.6Hz、1H)、7.88~7.75(d、J=5.9Hz、2H)、7.49~7.28(m、6H)、7.18(ddd、J=8.3、7.0、1.6Hz、1H)、6.88~6.75(m、2H)、6.49(dd、J=6.2、2.0Hz、1H)、6.16~6.02(m、2H)、5.93(s、2H)、5.05(d、J=3.7Hz、1H)、5.00~4.81(m、1H)、4.46(s、2H)、4.39~4.13(m、6H)、3.72~3.55(m、2H)、3.45~3.35(m、1H)、3.21(d、J=11.7Hz、2H)、2.98(d、J=11.6Hz、2H)、2.59(q、J=6.5Hz、4H)、2.42(s、6H)、2.17(dd、J=20.0、8.3Hz、3H)、1.94(s、4H)、1.75(t、J=9.6Hz、1H)、1.37(dd、J=20.9、7.0Hz、3H)、1.17(d、J=14.9Hz、1H)、1.05~0.87(m、3H)、0.77(dd、J=10.6、6.7Hz、3H)]として得た。 A 50 mL round-bottom flask was charged with a solution of (2S,4R)-1-[2-(3-[2-[4-(2-[[4-(3-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl]oxy]ethyl)piperazin-1-yl]ethoxy]-1,2-oxazol-5-yl)-3-methylbutanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (180 mg, 0.17 mmol, 1.0 equiv.) in methanol (5 mL) and hydrogen chloride (0.068 mL of a 4 M solution in dioxane, 0.27 mmol, 1.6 equiv.). The resulting solution was stirred at room temperature for 2 hours. The resulting mixture was concentrated under vacuum. The crude product was purified by chiral preparative HPLC using the following conditions: Column; Mobile phase, Column: CHIRALPAK ID-03, 2.0 cm I.D*25 cm L (5 um); Mobile phase A: MTBE (0.2% IPA)--HPLC, Mobile phase B: Methanol--HPLC, Flow rate: 18 mL/min, Gradient: 50B to 50B in 18 min, 254/220 nm. This gave 46.7 mg of (2S,4R)-1-[(2S)-2-[3-[2-(4-[2-[(4-[3-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl)oxy]ethyl]piperazin-1-yl)ethoxy]-1,2-oxazol-5-yl]-3-methylbutanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide as a white solid [1H NMR (300 MHz, DMSO-d6) δ 14.10 (s, 1H), 8.99-8.83 (m, 1H), 8.25 (d, J = 7.8 Hz, 1H), 7.92-7.83 (m, 2H), 7.75-7.12 (m, 6H), 6.81 (td, J = 8.4, 2.2 Hz, 2H), 6.49 (dd, J = 6.1, 2.0 Hz, 1H), 6.15-5.93 (m, 4H), 5.06 (d, J = 3.6H z, 1H), 4.98-4.79 (m, 1H), 4.38 (t, J = 7.7 Hz, 3H), 4.19 (dt, J = 21.0, 5.6 Hz, 5H), 3.98 (m, 1H), 3.71 (d, J = 8.6 Hz, 2H), 3.57-3.34 (m, 2H), 3.33-3.15 (m, 2H), 2.97 (d, J = 11.6 Hz, 4H), 2.57-2.40 (d, J = 8.0 Hz, 11H), 2.3 1-2.07 (m, 1H), 2.06-1.97 (m, 2H), 1.91-1.66 (m, 3H), 1.42 (d, J = 7.0 Hz, 1H), 1.31 (d, J = 7.0 Hz, 3H), 0.92-0.67 (m, 6H)], and 54.4 mg of (2S,4R)-1-[(2R)-2-[3-[2-(4-[2-[(4-[3-[3-amino-6-(2-hydroxyphenyl)pyridazinyl] ... [1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide as a white solid [1H NMR (300 MHz, DMSO-d6) δ 14.11 (s, 1H), 8.95 (s, 1H), 8.37 (d, J = 7.6 Hz, 1H), 7.88-7.75 (d, J = 5.9 Hz, 2H), 7.49-7.28 (m, 6H), 7.18 (ddd, J = 8.3, 7.0, 1.6 Hz, 1H), 6.88-6.75 (m, 2H), 6.49 (dd, J = 6.2, 2.0 Hz, 1H), 6.16-6.02 (m, 2H), 5.93 (s, 2H), 5.05 (d, J = 3.7 Hz, 1H), 5.00-4.81 (m, 1H), 4.46 (s, 2H), 4.39-4.13 (m , 6H), 3.72-3.55 (m, 2H), 3.45-3.35 (m, 1H), 3.21 (d, J = 11.7 Hz, 2H), 2.98 (d, J = 11.6 Hz, 2H), 2.59 (q, J = 6.5 Hz, 4H), 2.42 (s, 6H), 2.17 (dd, J = 20.0, 8.3 Hz, 3H), 1.94 (s, 4H), 1.75 (t, J = 9.6 Hz, 1H), 1.37 (dd, J = 20.9, 7.0 Hz, 3H), 1.17 (d, J = 14.9 Hz, 1H), 1.05-0.87 (m, 3H), 0.77 (dd, J = 10.6, 6.7 Hz, 3H)].
例示的な化合物48の例示的な合成
工程1
窒素不活性雰囲気下でパージし、維持した100mLの丸底フラスコに、メチル2-(6-クロロピリジン-3-イル)酢酸塩(1.86g、10.0mmol、1.00当量)、トルエン(50mL)、Cs2CO3(10.0g、30.0mmol、3.00当量)、1-ベンジルピペラジン(2.11g、12.0mmol、1.2当量)、およびRuPhosPd(0.39g、0.05当量)を入れた。得られた溶液を、油浴中で、100℃で10時間撹拌した。反応混合物を室温まで冷却した。次いで、反応を水(50mL)の添加により停止させた。得られた溶液を酢酸エチル(100mL×2)で抽出し、有機層を合わせた。得られた混合物をブラインで洗浄した。混合物を、無水硫酸ナトリウム上で乾燥させ、真空下で濃縮した。残渣を、酢酸エチル/石油エーテル(v:v=1/3)でシリカゲルカラム上に適用した。これにより、1.1g(32%)のメチル2-[6-(4-ベンジルピペラジン-1-イル)ピリジン-3-イル]酢酸塩を黄色の固体として得た。 A 100 mL round bottom flask purged and maintained under nitrogen inert atmosphere was charged with methyl 2-(6-chloropyridin-3-yl)acetate (1.86 g, 10.0 mmol, 1.00 equiv), toluene (50 mL), Cs2CO3 (10.0 g, 30.0 mmol, 3.00 equiv), 1-benzylpiperazine (2.11 g, 12.0 mmol, 1.2 equiv), and RuPhosPd (0.39 g, 0.05 equiv). The resulting solution was stirred in an oil bath at 100 °C for 10 h. The reaction mixture was cooled to room temperature. The reaction was then quenched by the addition of water (50 mL). The resulting solution was extracted with ethyl acetate (100 mL x 2) and the organic layers were combined. The resulting mixture was washed with brine. The mixture was dried over anhydrous sodium sulfate and concentrated under vacuum. The residue was applied onto a silica gel column with ethyl acetate/petroleum ether (v:v = 1/3). This yielded 1.1 g (32%) of methyl 2-[6-(4-benzylpiperazin-1-yl)pyridin-3-yl]acetate as a yellow solid.
工程2

窒素不活性雰囲気下でパージし、維持した100mLの三口丸底フラスコに、テトラヒドロフラン(30mL)、水素化アルミニウムリチウム(0.34g、3.00当量)を入れた。これに続いて、テトラヒドロフラン(5mL)中のメチル2-[6-(4-ベンジルピペラジン-1-イル)ピリジン-3-イル]酢酸塩(1.0g、3.01mmol、1.00当量)の溶液を、0℃で5分間撹拌しながら滴下で添加した。得られた溶液を、25℃で10時間撹拌した。次いで、反応物を、0℃で撹拌しながら、水(3mL)、水酸化ナトリウム水溶液(5%)、水(3mL)を添加することによってクエンチした。固体を濾過によって集めた。混合物を、無水硫酸ナトリウム上で乾燥させ、真空下で濃縮した。残渣を、ジクロロメタン/メタノール(v:v=1:10)でシリカゲルカラム上に適用した。これにより、750mg(81%)の2-[6-(4-ベンジルピペラジン-1-イル)ピリジン-3-イル]エタン-1-オールを淡い黄色の固体として得た。 A 100 mL three-necked round bottom flask purged and maintained under nitrogen inert atmosphere was charged with tetrahydrofuran (30 mL), lithium aluminum hydride (0.34 g, 3.00 equiv.). Following this, a solution of methyl 2-[6-(4-benzylpiperazin-1-yl)pyridin-3-yl]acetate (1.0 g, 3.01 mmol, 1.00 equiv.) in tetrahydrofuran (5 mL) was added dropwise with stirring at 0°C for 5 minutes. The resulting solution was stirred at 25°C for 10 hours. The reaction was then quenched by adding water (3 mL), aqueous sodium hydroxide (5%), water (3 mL) with stirring at 0°C. The solid was collected by filtration. The mixture was dried over anhydrous sodium sulfate and concentrated under vacuum. The residue was applied onto a silica gel column with dichloromethane/methanol (v:v=1:10). This yielded 750 mg (81%) of 2-[6-(4-benzylpiperazin-1-yl)pyridin-3-yl]ethan-1-ol as a pale yellow solid.
工程3

窒素不活性雰囲気下でパージし、維持した50mLの丸底フラスコに、2-[6-(4-ベンジルピペラジン-1-イル)ピリジン-3-イル]エタン-1-オール(750g、2.5mmol、1.00当量)、N,N-ジメチルホルムアミド(20mL)、水素化ナトリウム(60%)(200mg、8.4mmol、2.00当量)を0℃で入れた。得られた溶液を、0℃で0.5時間撹拌した。これに続いて、0℃で撹拌しながら、3-ブロモプロプ-1-イン(390mg、3.3mmol、1.30当量)を滴下で添加した。得られた溶液を、25℃でさらに12時間撹拌しながら反応させた。次いで、反応物を、0℃の水(20mL)を添加することによってクエンチした。得られた溶液を、酢酸エチル(20mL×2)で抽出し、有機層を合わせた。混合物を、無水硫酸ナトリウム上で乾燥させ、真空下で濃縮した。残渣を、酢酸エチル/石油エーテル(v:v=1:3)で溶出するシリカゲルカラム上に適用した。これにより、602mg(71%)の1-ベンジル-4-[5-[2-(プロパ-2-イン-1-イルオキシ)エチル]ピリジン-2-イル]ピペラジンを黄色の油として得た。 A 50 mL round bottom flask purged and maintained under nitrogen inert atmosphere was charged with 2-[6-(4-benzylpiperazin-1-yl)pyridin-3-yl]ethan-1-ol (750 g, 2.5 mmol, 1.00 equiv.), N,N-dimethylformamide (20 mL), sodium hydride (60%) (200 mg, 8.4 mmol, 2.00 equiv.) at 0°C. The resulting solution was stirred at 0°C for 0.5 h. Following this, 3-bromoprop-1-yne (390 mg, 3.3 mmol, 1.30 equiv.) was added dropwise with stirring at 0°C. The resulting solution was allowed to react with stirring at 25°C for an additional 12 h. The reaction was then quenched by adding water (20 mL) at 0°C. The resulting solution was extracted with ethyl acetate (20 mL x 2) and the organic layers were combined. The mixture was dried over anhydrous sodium sulfate and concentrated under vacuum. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (v:v=1:3). This gave 602 mg (71%) of 1-benzyl-4-[5-[2-(prop-2-yn-1-yloxy)ethyl]pyridin-2-yl]piperazine as a yellow oil.
工程4
窒素不活性雰囲気下でパージし、維持した25mLの密封チューブに、1-ベンジル-4-[5-[2-(プロパ-2-イン-1-イルオキシ)エチル]ピリジン-2-イル]ピペラジン(1.0g、3.00mmol、1.00当量)、テトラヒドロフラン(15mL)、ピナコールボラン(2mL)、ZrCp2HCl(0.077g、0.10当量)、トリエチルアミン(0.030g、0.10当量)を入れた。得られた溶液を、油浴中で、70℃で12時間撹拌した。次いで、反応物を、水(5mL)を添加することによってクエンチした。得られた溶液を酢酸エチル(10mL×2)で抽出し、有機層を合わせて、無水硫酸ナトリウムで乾燥させ、真空下で濃縮した。残渣を、ジクロロメタン/メタノール(v:v=10:1)で溶出するシリカゲルカラム上に適用した。これにより、1.05g(75%)の1-ベンジル-4-[5-(2-[[(2E)-3-(テトラメチル-1,3,2-ジオキサボロラン-2-イル)プロパ-2-エン-1-イル]オキシ]エチル)ピリジン-2-イル]ピペラジンをオレンジ色の油として得た。 In a 25 mL sealed tube purged and maintained under nitrogen inert atmosphere, 1-benzyl-4-[5-[2-(prop-2-yn-1-yloxy)ethyl]pyridin-2-yl]piperazine (1.0 g, 3.00 mmol, 1.00 equiv.), tetrahydrofuran (15 mL), pinacolborane (2 mL), ZrCp2HCl (0.077 g, 0.10 equiv.), triethylamine (0.030 g, 0.10 equiv.) were placed. The resulting solution was stirred in an oil bath at 70°C for 12 hours. The reaction was then quenched by adding water (5 mL). The resulting solution was extracted with ethyl acetate (10 mL x 2) and the organic layers were combined, dried over anhydrous sodium sulfate, and concentrated under vacuum. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (v:v=10:1). This gave 1.05 g (75%) of 1-benzyl-4-[5-(2-[[(2E)-3-(tetramethyl-1,3,2-dioxaborolan-2-yl)prop-2-en-1-yl]oxy]ethyl)pyridin-2-yl]piperazine as an orange oil.
工程5

100mLの丸底フラスコに、1-ベンジル-4-[5-(2-[[(2E)-3-(テトラメチル-1,3,2-ジオキサボロラン-2-イル)プロパ-2-エン-1-イル]オキシ]エチル)ピリジン-2-イル]ピペラジン(928mg、2.00mmol、1.00当量)、ジクロロメタン(25mL)を入れた。これに続いて、クロロギ酸1-クロロエチル(568.0mg、3.97mmol、2.00当量)を、0℃で撹拌しながら滴下で添加した。得られた溶液を、25℃で1時間撹拌した。得られた混合物を真空下で濃縮した。エチルアルコール(5mL)を添加した。温度を油浴中で80℃に維持しながら、得られた溶液をさらに2時間撹拌しながら反応させた。得られた混合物を真空下で濃縮した。これにより、0.71g(95%)の1-[5-(2-[[(2E)-3-(テトラメチル-1,3,2-ジオキサボロラン-2-イル)プロパ-2-エン-1-イル]オキシ]エチル)ピリジン-2-イル]ピペラジンを黄色の油として得た。 A 100 mL round bottom flask was charged with 1-benzyl-4-[5-(2-[[(2E)-3-(tetramethyl-1,3,2-dioxaborolan-2-yl)prop-2-en-1-yl]oxy]ethyl)pyridin-2-yl]piperazine (928 mg, 2.00 mmol, 1.00 equiv.), dichloromethane (25 mL). This was followed by the dropwise addition of 1-chloroethyl chloroformate (568.0 mg, 3.97 mmol, 2.00 equiv.) with stirring at 0°C. The resulting solution was stirred at 25°C for 1 hour. The resulting mixture was concentrated under vacuum. Ethyl alcohol (5 mL) was added. The resulting solution was allowed to react with stirring for an additional 2 hours while the temperature was maintained at 80°C in an oil bath. The resulting mixture was concentrated under vacuum. This gave 0.71 g (95%) of 1-[5-(2-[[(2E)-3-(tetramethyl-1,3,2-dioxaborolan-2-yl)prop-2-en-1-yl]oxy]ethyl)pyridin-2-yl]piperazine as a yellow oil.
工程6
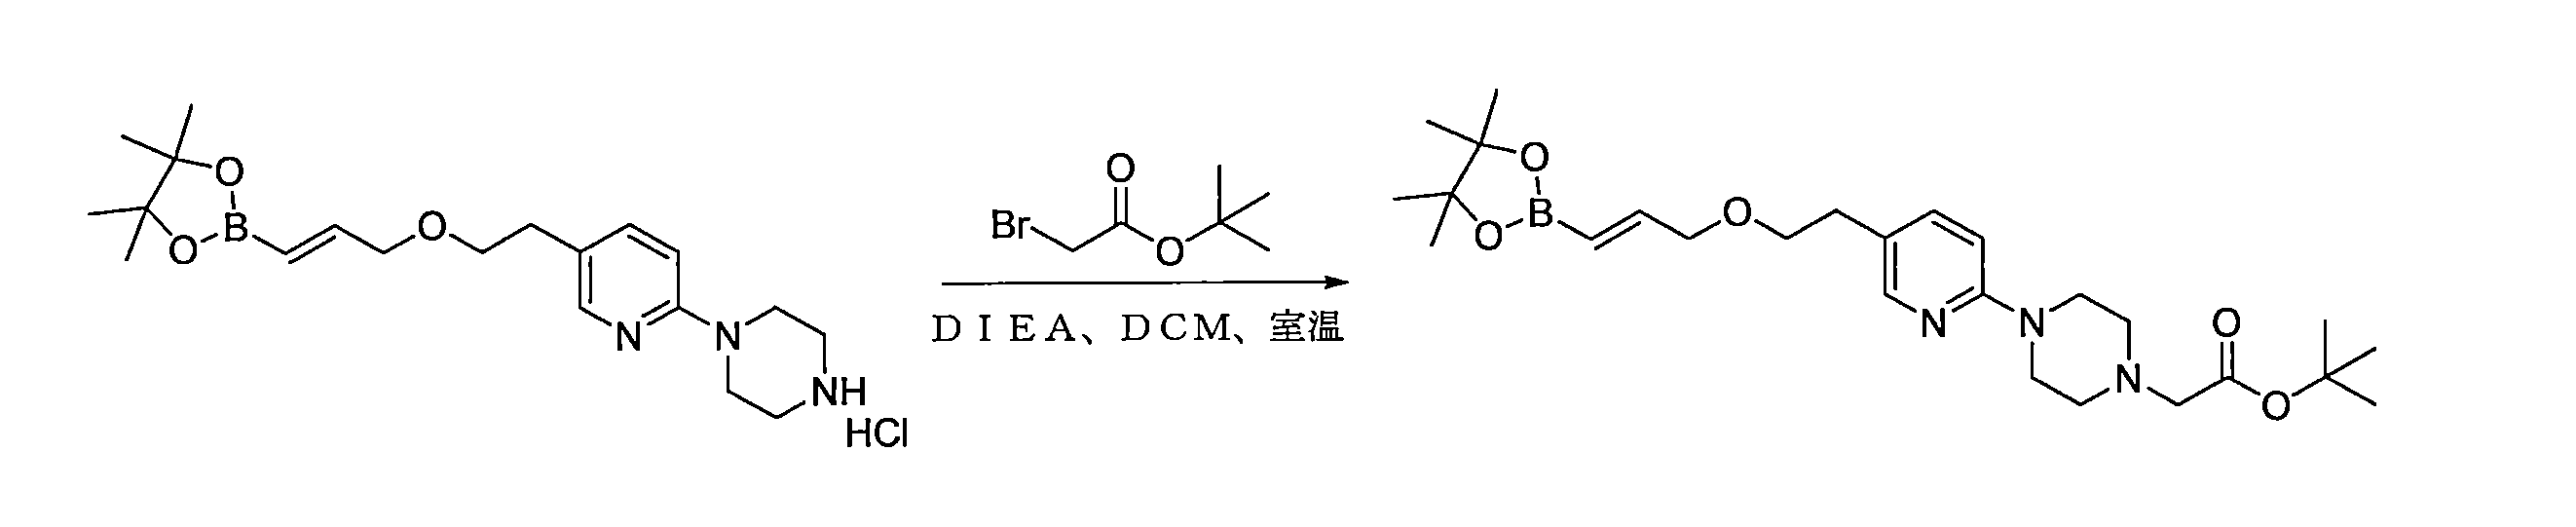
窒素不活性雰囲気下でパージし、維持した100mLの三口丸底フラスコに、ジクロロメタン中のN-(2-アミノエチル)-N-(2-クロロエチル)-5-(2-[[(2E)-3-(テトラメチル-1,3,2-ジオキサボロラン-2-イル)プロパ-2-エン-1-イル]オキシ]エチル)ピリジン-2-アミン(410mg、1.00mmol、1.00当量)を入れて、N,N-ジイソプロピルエチルアミン(645mg、4.99mmol、5.00当量)、およびtert-ブチル2-ブロモ酢酸塩(585mg、3.00mmol、3.00当量)を添加した。得られた溶液を25℃で12時間撹拌した。得られた混合物を、真空下で濃縮した。残渣を、ジクロロメタン/メタノール(v:v=10:1)で溶出するシリカゲルカラム上に適用した。これにより、0.32g(66%)のtert-ブチル2-[4-[5-(2-[[(2E)-3-(テトラメチル-1,3,2-ジオキサボロラン-2-イル)プロパ-2-エン-1-イル]オキシ]エチル)ピリジン-2-イル]ピペラジン-1-イル]酢酸塩を黄色の油として得た。 A 100 mL three-neck round bottom flask purged and maintained under nitrogen inert atmosphere was charged with N-(2-aminoethyl)-N-(2-chloroethyl)-5-(2-[[(2E)-3-(tetramethyl-1,3,2-dioxaborolan-2-yl)prop-2-en-1-yl]oxy]ethyl)pyridin-2-amine (410 mg, 1.00 mmol, 1.00 equiv.) in dichloromethane and N,N-diisopropylethylamine (645 mg, 4.99 mmol, 5.00 equiv.), and tert-butyl 2-bromoacetate (585 mg, 3.00 mmol, 3.00 equiv.). The resulting solution was stirred at 25°C for 12 hours. The resulting mixture was concentrated under vacuum. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (v:v=10:1). This yielded 0.32 g (66%) of tert-butyl 2-[4-[5-(2-[[(2E)-3-(tetramethyl-1,3,2-dioxaborolan-2-yl)prop-2-en-1-yl]oxy]ethyl)pyridin-2-yl]piperazin-1-yl]acetate as a yellow oil.
工程7

窒素不活性雰囲気下でパージし、維持した25mLの密封チューブに、ジオキサン/水(v:v=4:1)中の2-6-アミノ-5-[8-(2-ブロモピリジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-3-イル]ピリダジン-3-イルフェノール(158mg、0.35mmol、1.00当量)を入れて、tert-ブチル2-[4-[5-(2-[[(2E)-3-(テトラメチル-1,3,2-ジオキサボロラン-2-イル)プロパ-2-エン-1-イル]オキシ]エチル)ピリジン-2-イル]ピペラジン-1-イル]酢酸塩(205mg、0.42mmol、1.20当量)、炭酸カリウム(145mg、1.05mmol、3.00当量)、およびPd(dppf)Cl2(12mg、0.02mmol、0.05当量)を添加した。得られた溶液を、油浴中で、80℃で3時間撹拌した。反応混合物を、25℃に冷却した。次いで、反応物を、水(5mL)を添加することによってクエンチした。得られた溶液を酢酸エチル(20mL×2)で抽出し、有機層を合わせて、無水硫酸ナトリウムで乾燥させ、真空下で濃縮した。残渣を、ジクロロメタン/メタノール(v:v=10:1)でシリカゲルカラム上に適用した。これにより、0.16g(63%)のtert-ブチル2-[4-[5-(2-[[(2E)-3-(4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)プロパ-2-エン-1-イル]オキシ]エチル)ピリジン-2-イル]ピペラジン-1-イル]酢酸塩を淡い黄色の固体として得た。 In a 25 mL sealed tube purged and maintained under a nitrogen inert atmosphere, 2-6-amino-5-[8-(2-bromopyridin-4-yl)-3,8-diazabicyclo[3.2.1]octan-3-yl]pyridazin-3-ylphenol (158 mg, 0.35 mmol, 1.00 equiv.) in dioxane/water (v:v=4:1) was added to prepare tert-butyl 2-[4-[5-(2-[[(2 E)-3-(tetramethyl-1,3,2-dioxaborolan-2-yl)prop-2-en-1-yl]oxy]ethyl)pyridin-2-yl]piperazin-1-yl]acetate (205 mg, 0.42 mmol, 1.20 equiv.), potassium carbonate (145 mg, 1.05 mmol, 3.00 equiv.), and Pd(dppf)Cl2 (12 mg, 0.02 mmol, 0.05 equiv.) were added. The resulting solution was stirred at 80° C. in an oil bath for 3 hours. The reaction mixture was cooled to 25° C. Then, the reaction was quenched by adding water (5 mL). The resulting solution was extracted with ethyl acetate (20 mL×2) and the organic layers were combined, dried over anhydrous sodium sulfate, and concentrated under vacuum. The residue was applied onto a silica gel column with dichloromethane/methanol (v:v=10:1). This yielded 0.16 g (63%) of tert-butyl 2-[4-[5-(2-[[(2E)-3-(4-[3-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl)prop-2-en-1-yl]oxy]ethyl)pyridin-2-yl]piperazin-1-yl]acetate as a pale yellow solid.
他の実施例について上に記載される手順、ならびに当業者によって理解される手順を使用して、以下のスキームに従って、tert-ブチル2-[4-[5-(2-[[(2E)-3-(4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)プロパ-2-エン-1-イル]オキシ]エチル)ピリジン-2-イル]ピペラジン-1-イル]酢酸塩を、最終化合物(2S,4R)-1-((2S)-2-(2-(4-(5-(2-(3-(4-(3-(3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル)プロポキシ)エチル)ピリジン-2-イル)ピペラジン-1-イル)アセトアミド)-3,3-ジメチルブタノイル)-4-ヒドロキシ-N-(4-(4-メチルチアゾール-5-イル)ベンジル)ピロリジン-2-カルボキサミドに変換した。

例示的な化合物49の例示的な合成
工程1
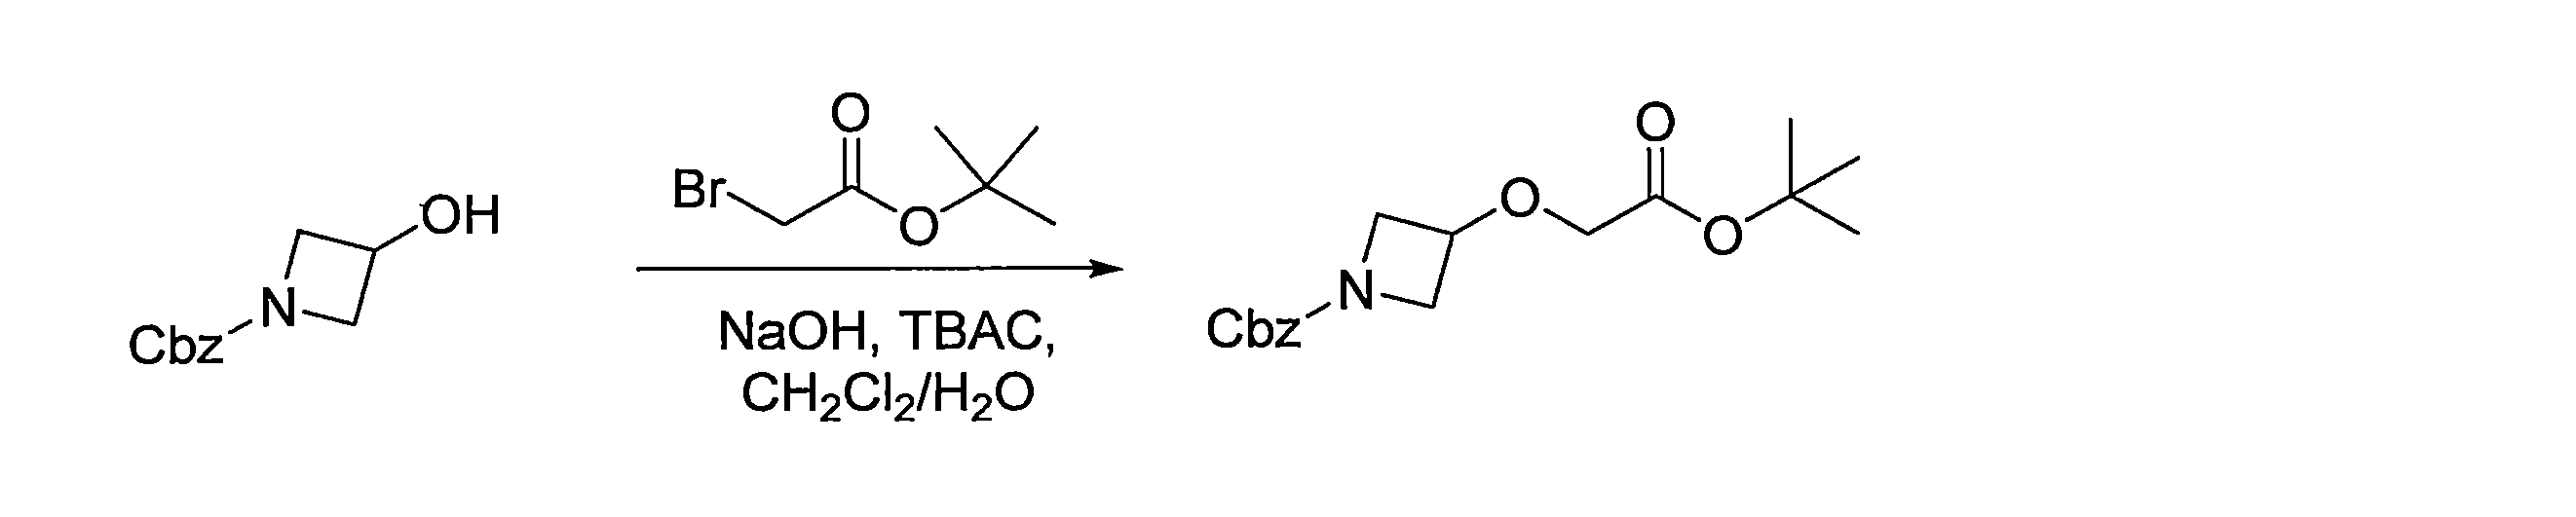
100mLの丸底フラスコに、ベンジル3-ヒドロキシアゼチジン-1-カルボキシレート(3.11g、15.01mmol、1.00当量)、ジクロロメタン(30mL)、2-ブロモ酢酸tert-ブチル(4.37g、22.40mmol、1.50当量)、水酸化ナトリウム(37%水溶液)(30mL)、およびTBAC(4.17g、1.00当量)を0℃で入れた。得られた溶液を、25℃で3時間撹拌した。得られた溶液をジクロロメタン(2×30mL)で抽出し、有機層を合わせ、無水硫酸ナトリウム上で乾燥させ、真空下で濃縮した。残渣を、酢酸エチル/石油エーテル(1/5)で溶出するシリカゲルカラム上に適用した。これにより、3.6g(75%)のベンジル3-[2-(tert-ブトキシ)-2-オキソエトキシ]アゼチジン-1-カルボキシレートを無色の油として得た。 A 100 mL round bottom flask was charged with benzyl 3-hydroxyazetidine-1-carboxylate (3.11 g, 15.01 mmol, 1.00 equiv.), dichloromethane (30 mL), tert-butyl 2-bromoacetate (4.37 g, 22.40 mmol, 1.50 equiv.), sodium hydroxide (37% in water) (30 mL), and TBAC (4.17 g, 1.00 equiv.) at 0°C. The resulting solution was stirred at 25°C for 3 hours. The resulting solution was extracted with dichloromethane (2 x 30 mL), the organic layers were combined, dried over anhydrous sodium sulfate, and concentrated under vacuum. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (1/5). This gave 3.6 g (75%) of benzyl 3-[2-(tert-butoxy)-2-oxoethoxy]azetidine-1-carboxylate as a colorless oil.
工程2

250mLの丸底フラスコに、ベンジル3-[2-(tert-ブトキシ)-2-オキソエトキシ]アゼチジン-1-カルボキシレート(3.22g、10.02mmol、1.00当量)、エタノール(100mL)、炭素上のパラジウム(0.322g)を入れた。得られた溶液を25℃においてH2雰囲気下で12時間撹拌した。固体を濾過によって集めた。得られた混合物を真空下で濃縮した。これにより、1.75g(93%)のtert-ブチル2-(アゼチジン-3-イルオキシ)酢酸塩を無色の油として得た。 A 250 mL round bottom flask was charged with benzyl 3-[2-(tert-butoxy)-2-oxoethoxy]azetidine-1-carboxylate (3.22 g, 10.02 mmol, 1.00 equiv), ethanol (100 mL), and palladium on carbon (0.322 g). The resulting solution was stirred at 25°C under an H2 atmosphere for 12 h. The solids were collected by filtration. The resulting mixture was concentrated under vacuum. This gave 1.75 g (93%) of tert-butyl 2-(azetidin-3-yloxy)acetate as a colorless oil.
工程3

50mLの丸底フラスコに、tert-ブチル2-(アゼチジン-3-イルオキシ)酢酸塩(374mg、2.00mmol、1.00当量)、N,N-ジメチルホルムアミド(15mL)、炭酸カリウム(828mg、5.99mmol、3.00当量)、および1,2-ジブロモエタン(1.86g、9.90mmol、5.00当量)を入れた。得られた溶液を25℃で12時間撹拌した。次いで、反応物を、水(10mL)を添加することによってクエンチした。得られた溶液を、酢酸エチル(2×20mL)で抽出し、有機層を合わせて、無水硫酸ナトリウムで乾燥させた。残渣を、酢酸エチル/石油エーテル(1/3)で溶出するシリカゲルカラム上に適用した。これにより、80mg(14%)のtert-ブチル2-[[1-(2-ブロモエチル)アゼチジン-3-イル]オキシ]酢酸塩を無色の油として得た。 A 50 mL round bottom flask was charged with tert-butyl 2-(azetidin-3-yloxy)acetate (374 mg, 2.00 mmol, 1.00 equiv), N,N-dimethylformamide (15 mL), potassium carbonate (828 mg, 5.99 mmol, 3.00 equiv), and 1,2-dibromoethane (1.86 g, 9.90 mmol, 5.00 equiv). The resulting solution was stirred at 25°C for 12 hours. The reaction was then quenched by adding water (10 mL). The resulting solution was extracted with ethyl acetate (2 x 20 mL) and the organic layers were combined and dried over anhydrous sodium sulfate. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (1/3). This gave 80 mg (14%) of tert-butyl 2-[[1-(2-bromoethyl)azetidin-3-yl]oxy]acetate as a colorless oil.
他の実施例について上に記載される手順、ならびに当業者に明らかである手順を使用して、tert-ブチル2-[[1-(2-ブロモエチル)アゼチジン-3-イル]オキシ]酢酸塩を、最終化合物(2S,4R)-1-((2S)-2-(2-((1-(2-(4-(3-(4-(3-(3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル)プロピル)ピペラジン-1-イル)エチル)アゼチジン-3-イル)オキシ)アセトアミド)-3,3-ジメチルブタノイル)-4-ヒドロキシ-N-((S)-1-(4-(4-メチルチアゾール-5-イル)フェニル)エチル)ピロリジン-2-カルボキサミドに変換した。

例示的な化合物51の例示的な合成
例示的な化合物51を、上に記載される実施例(例えば、例示的な化合物37)からの手順、および当業者によって理解される手順を使用して、以下のスキームに従って調製した。

例示的な化合物52の例示的な合成
例示的な化合物52を、以下のスキームに従って調製した。

例示的な化合物53の例示的な合成
例示的な化合物53を、以下のスキームに従って調製した。

例示的な化合物54および例示的な化合物55の例示的な合成
工程1

100mLの丸底フラスコに、2-(3-メトキシ-1,2-オキサゾール-5-イル)-3-メチルブタン酸(2.1g、10.5mmol、1.00当量)、エタノール(20.0g、434mmol、41.2当量)、硫酸(2.0g、20.4mmol、1.9当量)を入れた。得られた溶液を、油浴中で、70℃で12時間撹拌した。得られた混合物を真空下で濃縮した。これにより、1.9g(79%)のエチル2-(3-メトキシ-1,2-オキサゾール-5-イル)-3-メチルブタノエートを白色の固体として得た。 A 100 mL round bottom flask was charged with 2-(3-methoxy-1,2-oxazol-5-yl)-3-methylbutanoic acid (2.1 g, 10.5 mmol, 1.00 equiv.), ethanol (20.0 g, 434 mmol, 41.2 equiv.), and sulfuric acid (2.0 g, 20.4 mmol, 1.9 equiv.). The resulting solution was stirred in an oil bath at 70° C. for 12 hours. The resulting mixture was concentrated under vacuum. This afforded 1.9 g (79%) of ethyl 2-(3-methoxy-1,2-oxazol-5-yl)-3-methylbutanoate as a white solid.
工程2

50mLの丸底フラスコに、エチル2-(3-メトキシ-1,2-オキサゾール-5-イル)-3-メチルブタノエート(1.9g、8.4mmol、1.0当量)および塩化スルフリル(20mL)を入れた。得られた溶液を、室温で一晩撹拌した。得られた混合物を真空下で濃縮した。次いで、反応を30mLの水の添加によりクエンチさせた。得られた溶液を酢酸エチル(30mL×3)で抽出し、有機層を合わせた。残渣を、酢酸エチル/石油エーテル(1:1)で溶出するシリカゲルカラム上に適用した。これにより、1.5g(69%)のエチル2-(4-クロロ-3-メトキシ-1、2-オキサゾール-5-イル)-3-メチルブタノエートを黄色の油として得た。 A 50 mL round bottom flask was charged with ethyl 2-(3-methoxy-1,2-oxazol-5-yl)-3-methylbutanoate (1.9 g, 8.4 mmol, 1.0 equiv.) and sulfuryl chloride (20 mL). The resulting solution was stirred at room temperature overnight. The resulting mixture was concentrated under vacuum. The reaction was then quenched by the addition of 30 mL of water. The resulting solution was extracted with ethyl acetate (30 mL x 3) and the organic layers were combined. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (1:1). This gave 1.5 g (69%) of ethyl 2-(4-chloro-3-methoxy-1,2-oxazol-5-yl)-3-methylbutanoate as a yellow oil.
工程3
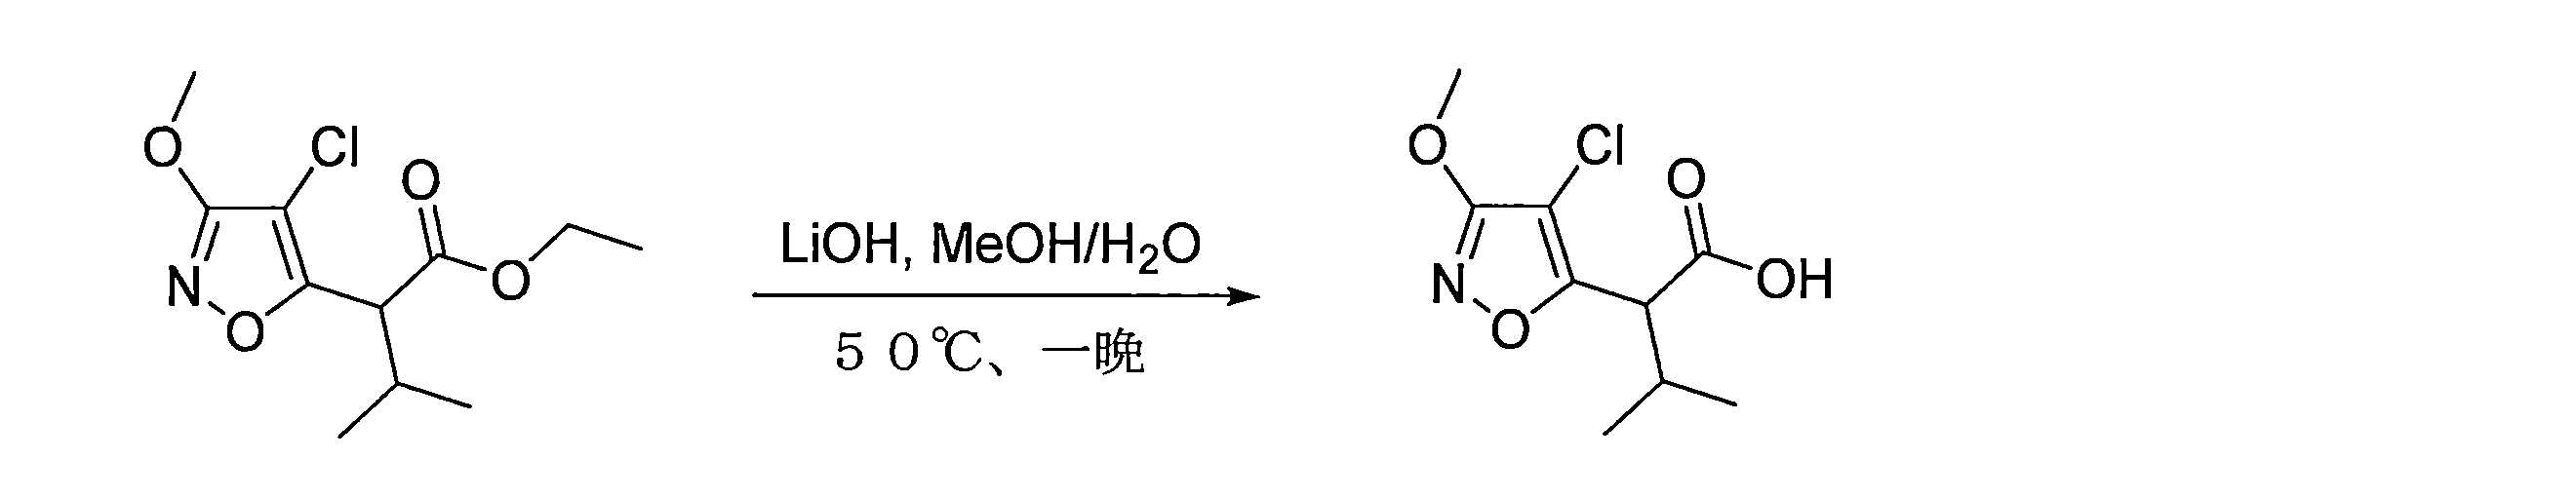
50mLの丸底フラスコに、メタノール/H2O(5/5mL)中のエチル2-(4-クロロ-3-メトキシ-1,2-オキサゾール-5-イル)-3-メチルブタノエート(1.5g、5.7mmol、1.0当量)の溶液、および水酸化リチウム(300mg、12.5mmol、2.2当量)を入れた。得られた溶液を、油浴中で、50℃で一晩撹拌した。溶液のpH値を、1M HClで6に調整した。得られた溶液を酢酸エチル(30mL×3)で抽出し、有機層を合わせて、真空下で濃縮した。以下の条件の分取HPLCによって、粗生成物を精製した:カラム;移動相、カラム:XBridge Prep C18 OBD Column、5um、19*150mm;移動相A:水(0.1%FA)、移動相B:ACN、流速:20mL/分、勾配:8分間で34%B~48%B、220nm。これにより、300mg(22.4%)の2-(4-クロロ-3-メトキシ-1,2-オキサゾール-5-イル)-3-メチルブタン酸を白色の固体として得た。 A 50 mL round bottom flask was charged with a solution of ethyl 2-(4-chloro-3-methoxy-1,2-oxazol-5-yl)-3-methylbutanoate (1.5 g, 5.7 mmol, 1.0 equiv.) in methanol/H2O (5/5 mL) and lithium hydroxide (300 mg, 12.5 mmol, 2.2 equiv.). The resulting solution was stirred overnight at 50 °C in an oil bath. The pH value of the solution was adjusted to 6 with 1 M HCl. The resulting solution was extracted with ethyl acetate (30 mL x 3) and the organic layers were combined and concentrated under vacuum. The crude product was purified by preparative HPLC with the following conditions: Column; Mobile phase, Column: XBridge Prep C18 OBD Column, 5um, 19*150mm; Mobile phase A: Water (0.1% FA), Mobile phase B: ACN, Flow rate: 20mL/min, Gradient: 34%B to 48%B in 8min, 220nm. This gave 300mg (22.4%) of 2-(4-chloro-3-methoxy-1,2-oxazol-5-yl)-3-methylbutanoic acid as a white solid.
工程4
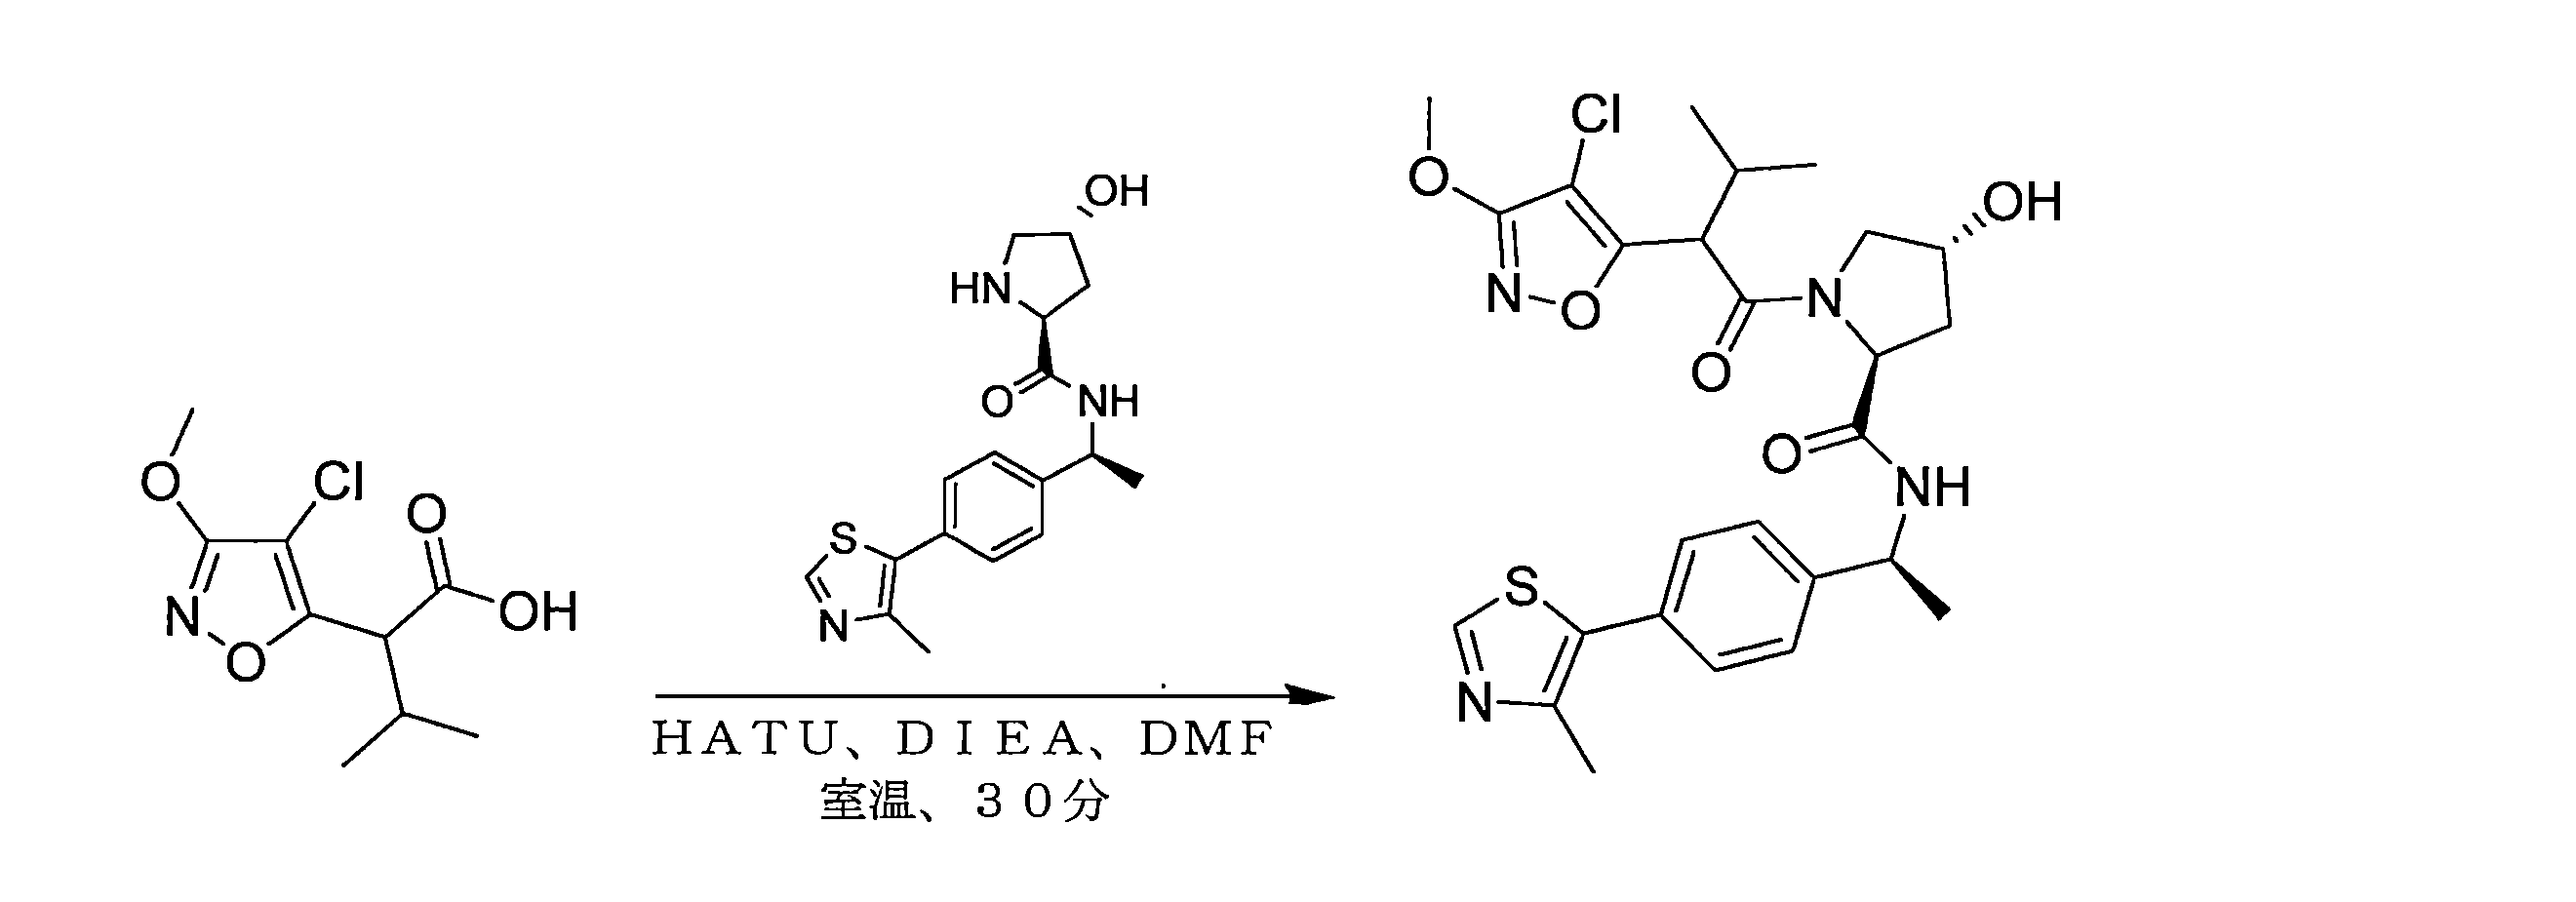
50mLの丸底フラスコに、N,N-ジメチルホルムアミド(20mL)中の2-(4-クロロ-3-メトキシ-1,2-オキサゾール-5-イル)-3-メチルブタン酸(260mg、1.1mmol、1.0当量)の溶液、N,N,N’,N’-テトラメチル-O-(7-アザベンゾトリアゾール-1-イル)ウロニウムヘキサフルオロホスフェート(360mg、1.0mmol、0.9当量)、N,N-ジイソプロピルエチルアミン(350mg、2.7mmol、2.4当量)、および(2S,4R)-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(350mg、1.1mmol、1.0当量)を入れた。得られた溶液を室温で30分間撹拌した。次いで、反応を30mLの水の添加によりクエンチさせた。得られた溶液を酢酸エチル(30mL×3)で抽出し、有機層を合わせて、無水硫酸ナトリウムで乾燥させ、真空下で濃縮した。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。これにより、275mg(45%)の(2S,4R)-1-[2-(4-クロロ-3-メトキシ-1,2-オキサゾール-5-イル)-3-メチルブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミドを白色の固体として得た。 A 50 mL round-bottom flask was charged with a solution of 2-(4-chloro-3-methoxy-1,2-oxazol-5-yl)-3-methylbutanoic acid (260 mg, 1.1 mmol, 1.0 equiv.) in N,N-dimethylformamide (20 mL), N,N,N',N'-tetramethyl-O-(7-azabenzotriazol-1-yl)uronium hexafluorophosphate (360 mg, 1.0 mmol, 0.9 equiv.), N,N-diisopropylethylamine (350 mg, 2.7 mmol, 2.4 equiv.), and (2S,4R)-4-hydroxy-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (350 mg, 1.1 mmol, 1.0 equiv.). The resulting solution was stirred at room temperature for 30 min. The reaction was then quenched by the addition of 30 mL of water. The resulting solution was extracted with ethyl acetate (30 mL x 3) and the organic layers were combined, dried over anhydrous sodium sulfate and concentrated under vacuum. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (10:1). This gave 275 mg (45%) of (2S,4R)-1-[2-(4-chloro-3-methoxy-1,2-oxazol-5-yl)-3-methylbutanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide as a white solid.
工程5
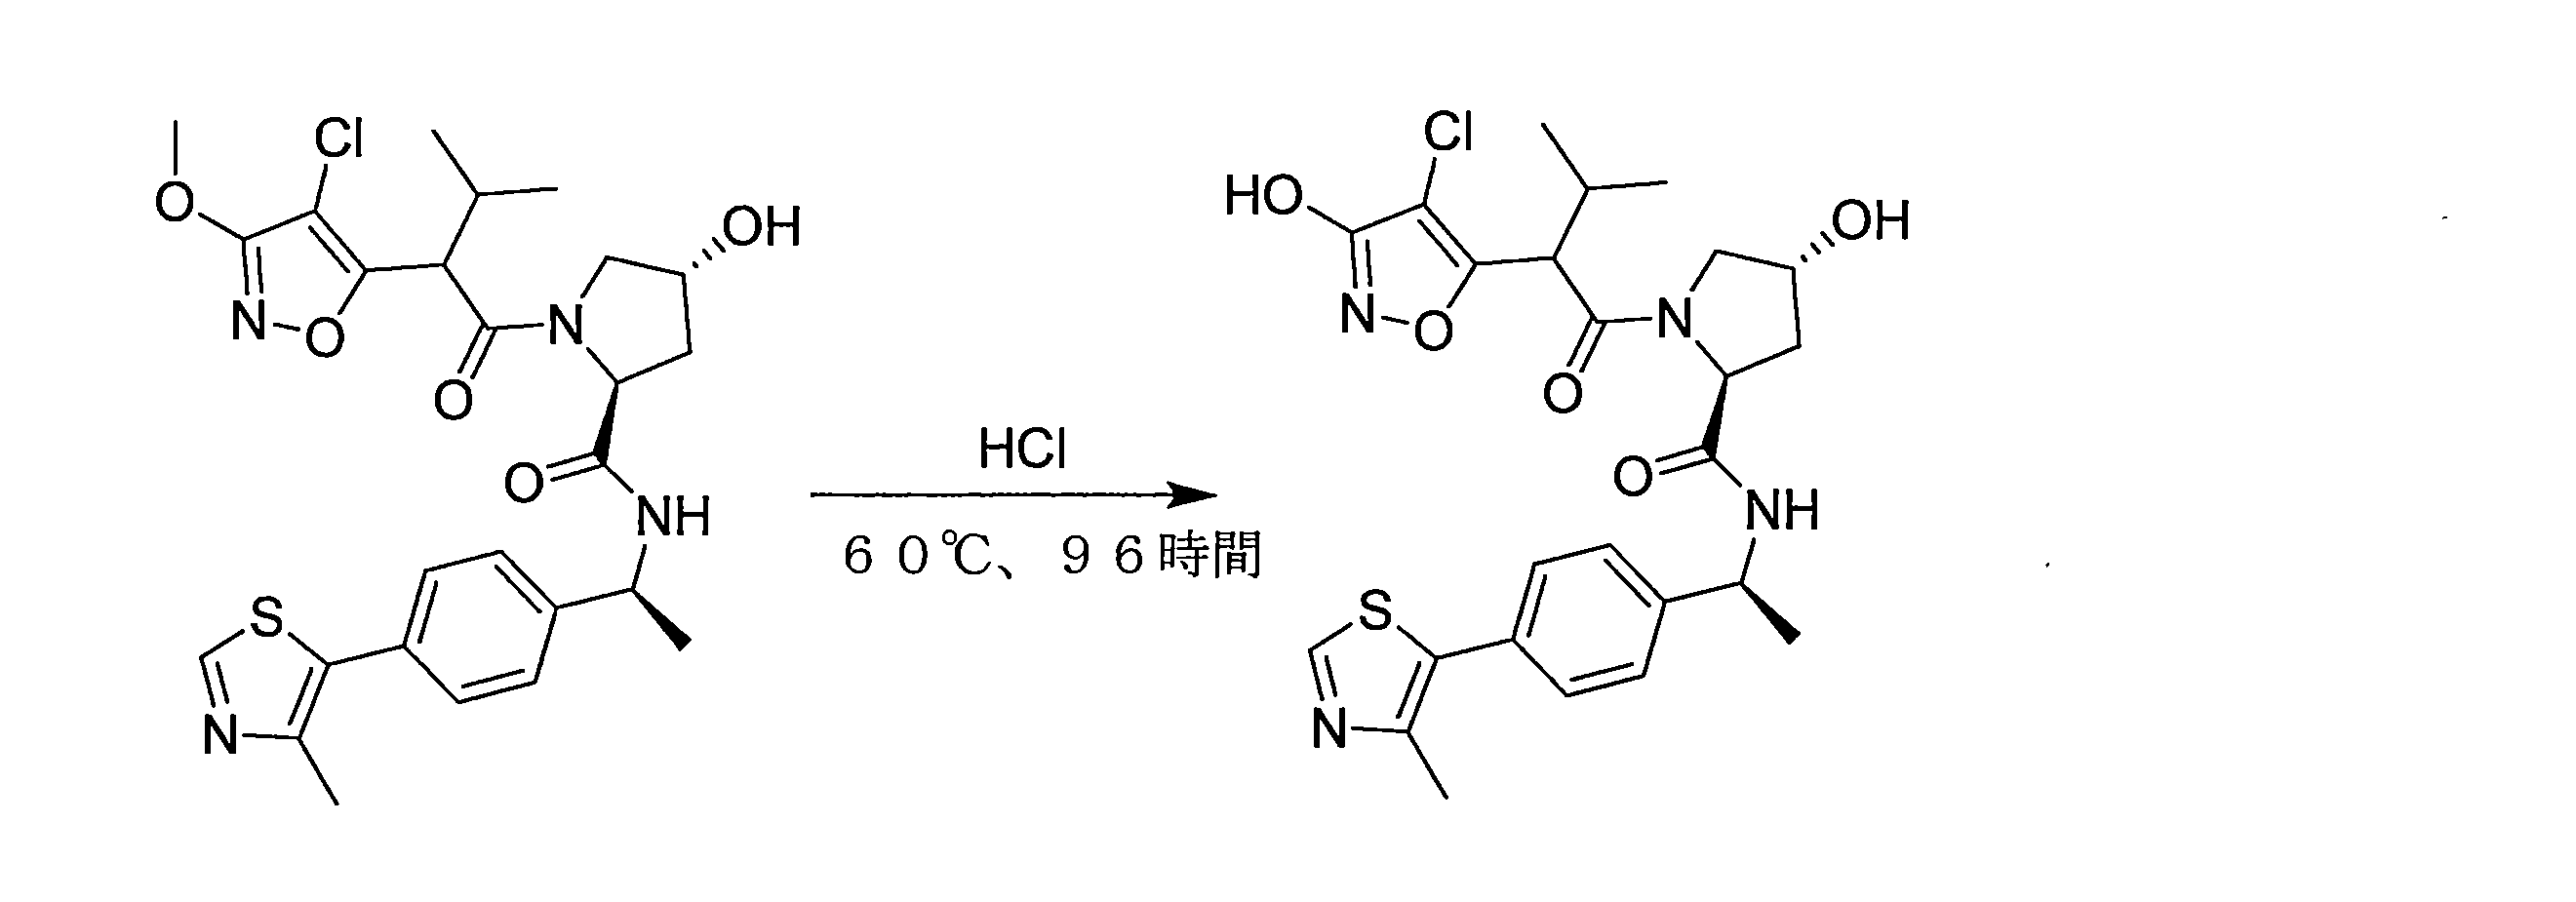
50mLの丸底フラスコに、(2S,4R)-1-[2-(4-クロロ-3-メトキシ-1,2-オキサゾール-5-イル)-3-メチルブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(275mg、0.5mmol、1.0当量)およびジオキサン(25mL)中の4M塩化水素溶液を入れた。得られた溶液を、油浴中で、60℃で96時間撹拌した。得られた混合物を真空下で濃縮した。これにより、259mg(97%)の(2S,4R)-1-[2-(4-クロロ-3-ヒドロキシ-1,2-オキサゾール-5-イル)-3-メチルブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミドを黄色の固体として得た。 A 50 mL round bottom flask was charged with (2S,4R)-1-[2-(4-chloro-3-methoxy-1,2-oxazol-5-yl)-3-methylbutanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (275 mg, 0.5 mmol, 1.0 equiv.) and a 4 M solution of hydrogen chloride in dioxane (25 mL). The resulting solution was stirred in an oil bath at 60 °C for 96 h. The resulting mixture was concentrated under vacuum. This gave 259 mg (97%) of (2S,4R)-1-[2-(4-chloro-3-hydroxy-1,2-oxazol-5-yl)-3-methylbutanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide as a yellow solid.
以下のスキームに従って、かつ上記の前述の実施例について記載される手順を使用して、(2S,4R)-1-[2-(4-クロロ-3-ヒドロキシ-1,2-オキサゾール-5-イル)-3-メチルブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミドを、最終化合物(2S,4R)-1-((2S)-2-(3-(2-(4-(2-((4-(3-(3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル)オキシ)エチル)ピペラジン-1-イル)エトキシ)-4-クロロイソキサゾール-5-イル)-3-メチルブタノイル)-4-ヒドロキシ-N-((S)-1-(4-(4-メチルチアゾール-5-イル)フェニル)エチル)ピロリジン-2-カルボキサミドおよび(2S,4R)-1-((2R)-2-(3-(2-(4-(2-((4-(3-(3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル)オキシ)エチル)ピペラジン-1-イル)エトキシ)-4-クロロイソキサゾール-5-イル)-3-メチルブタノイル)-4-ヒドロキシ-N-((S)-1-(4-(4-メチルチアゾール-5-イル)フェニル)エチル)ピロリジン-2-カルボキサミドに変換した。
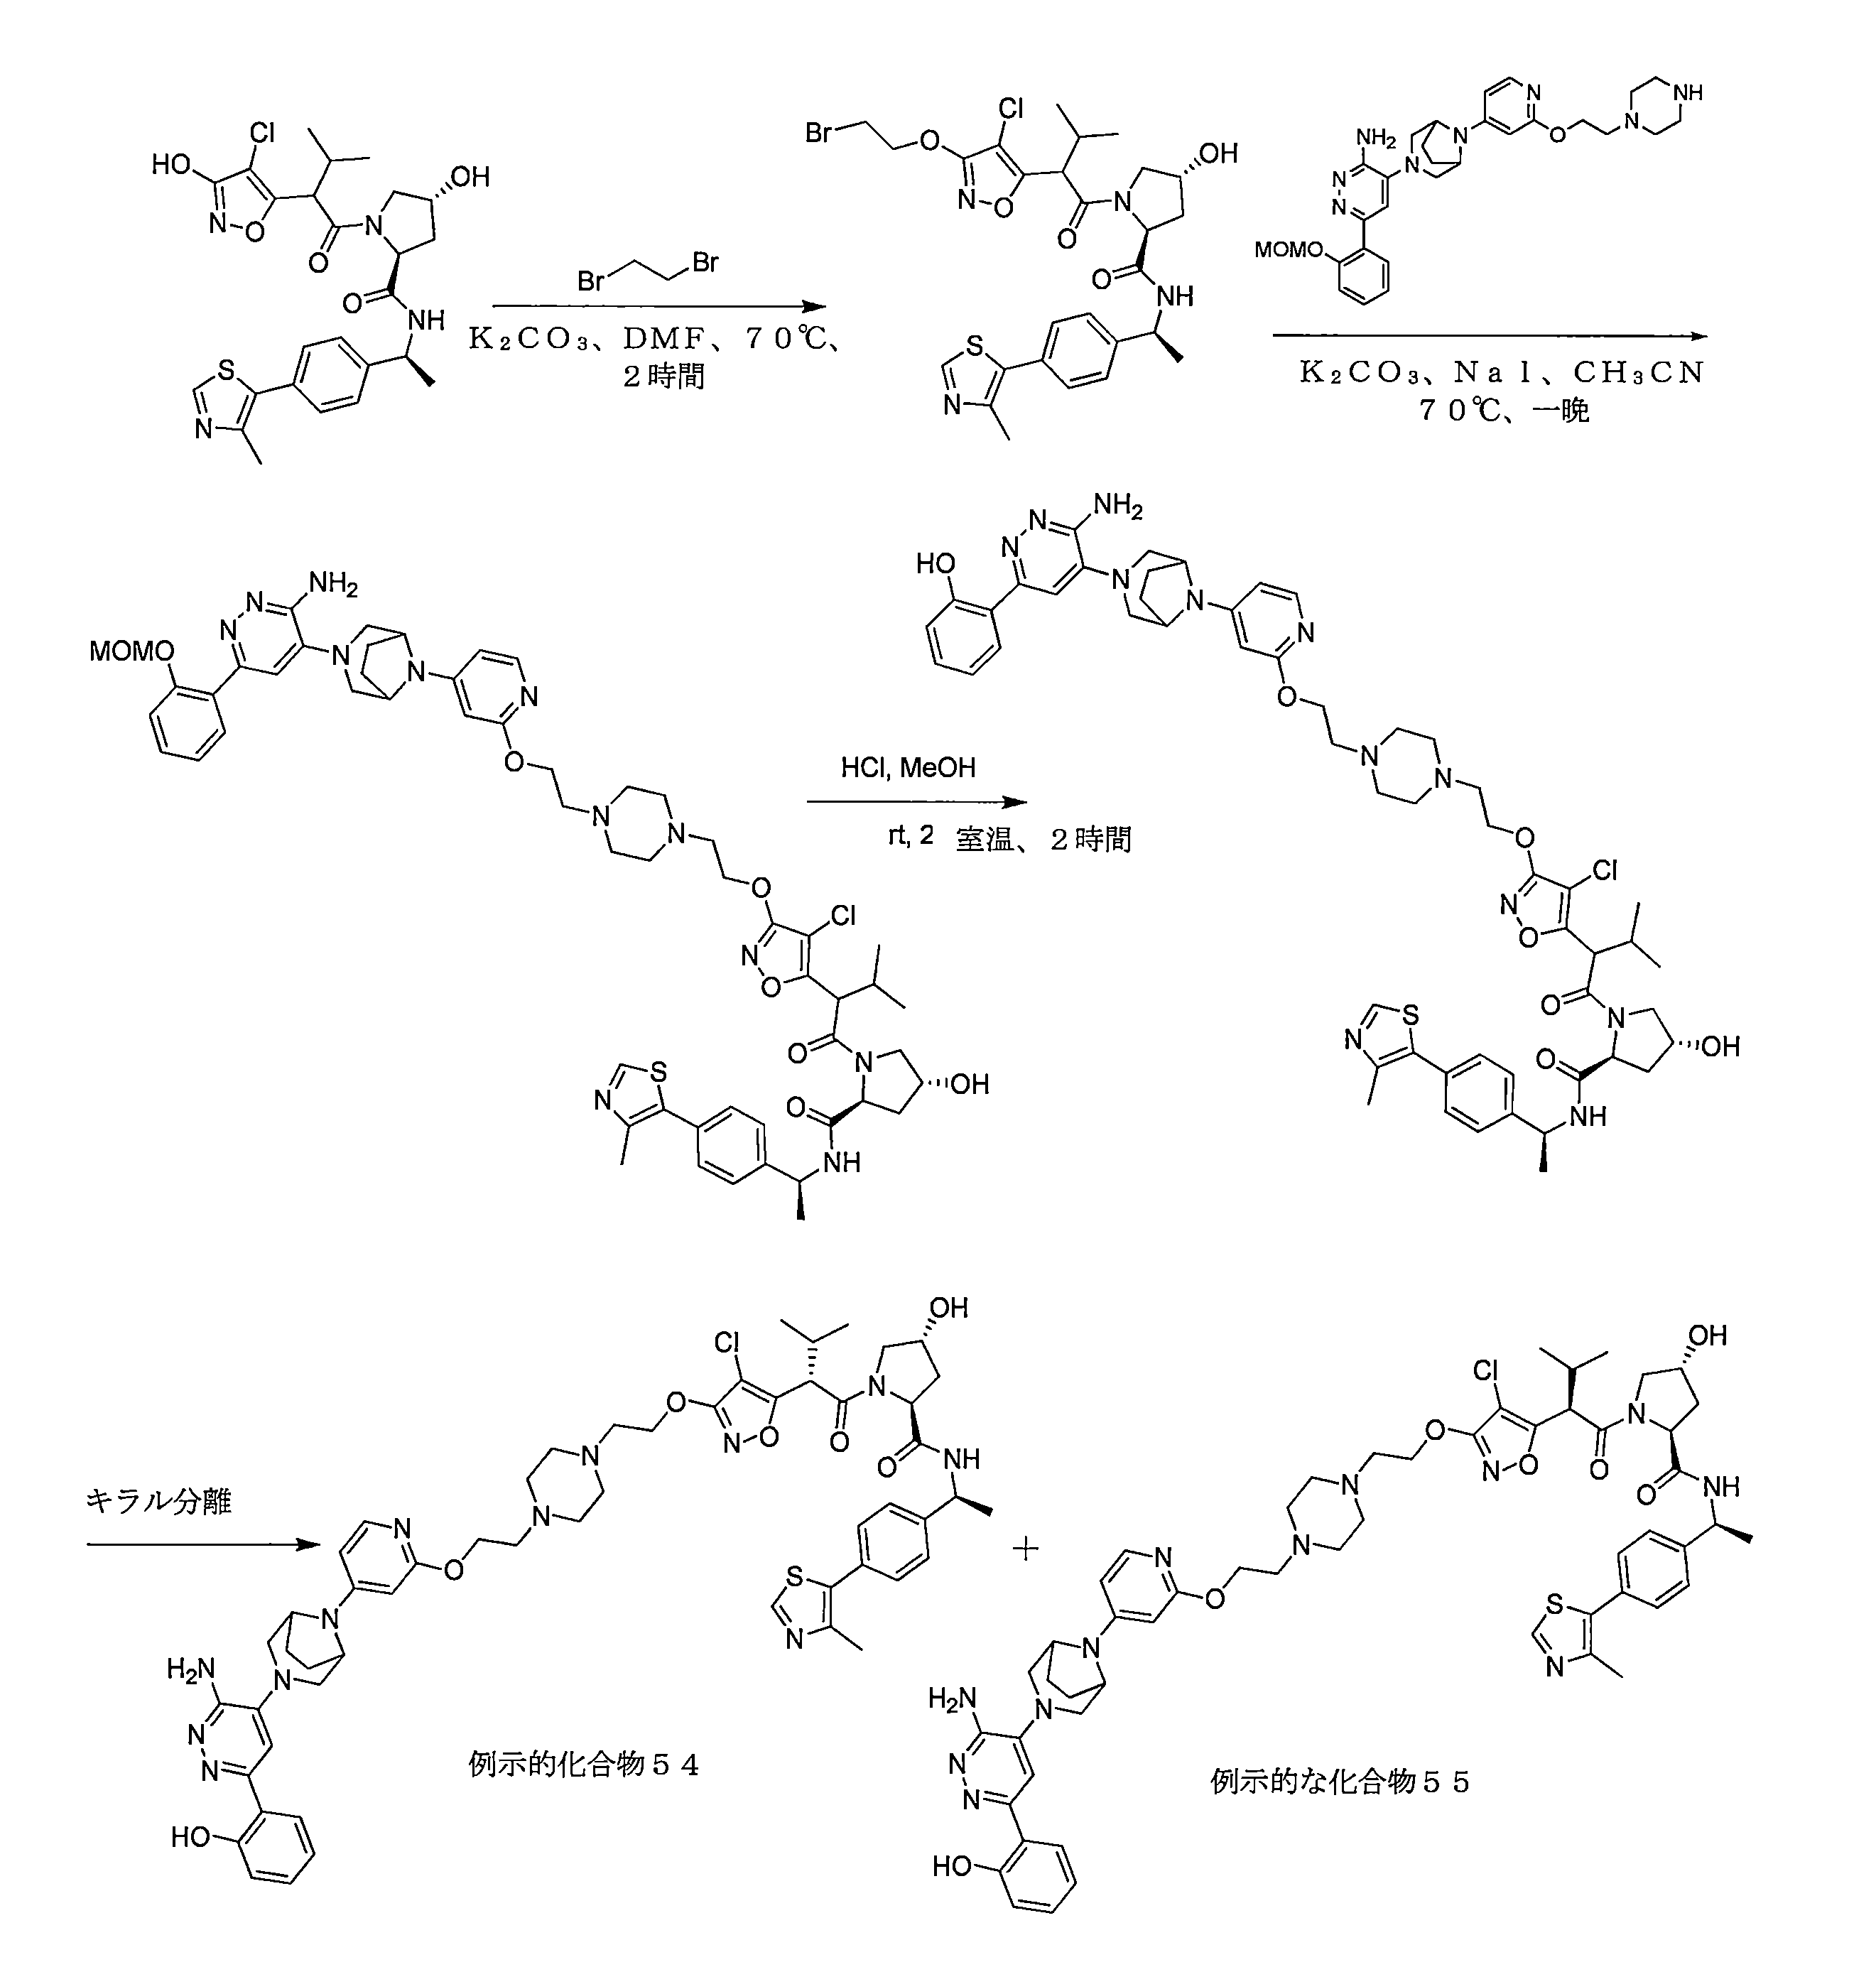
例示的な化合物56の例示的な合成
工程1
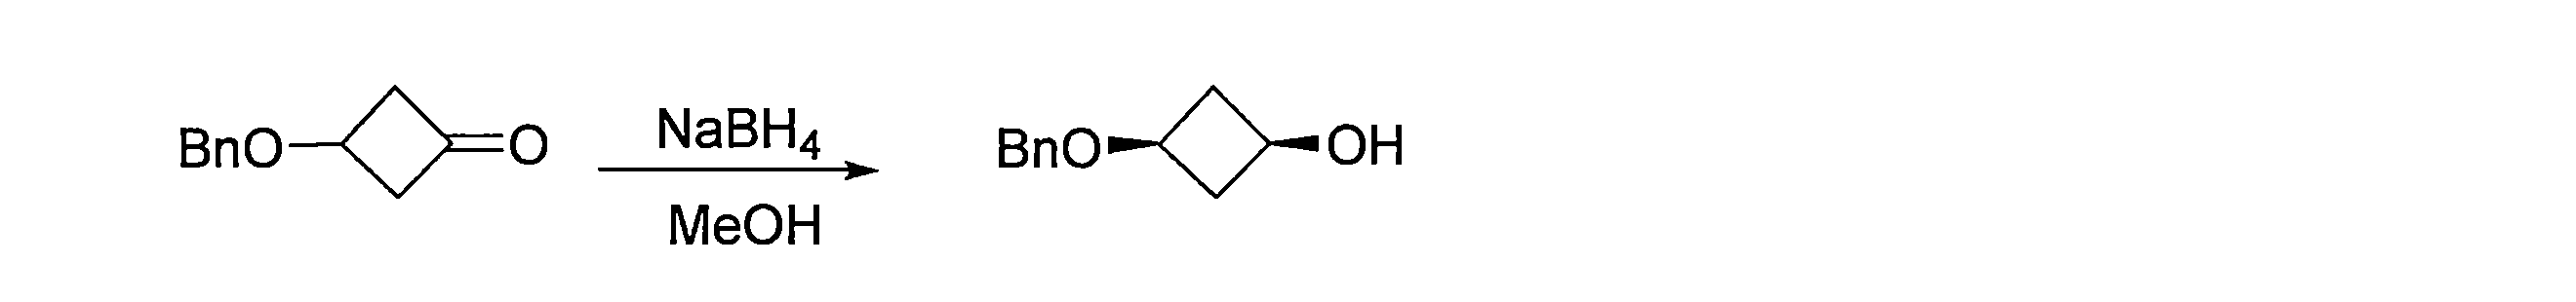
250mLの丸底フラスコに、エタノール(100mL)中の3-(ベンジルオキシ)シクロブタン-1-オン(6.0g、34.05mmol、1.0当量)の溶液を入れた。これに続いて、0℃で、水素化ホウ素ナトリウム(1.25g、33.94mmol、1.00当量)を数回に分けて添加した。得られた溶液を、0℃で3時間撹拌した。次いで、反応物を、水(100mL)を添加することによってクエンチした。得られた溶液を酢酸エチル(50mL×2)で抽出し、有機層を合わせて、無水硫酸ナトリウムで乾燥させ、真空下で濃縮した。これにより、6.0g(99%)の(1s,3s)-3-(ベンジルオキシ)シクロブタン-1-オールを淡い黄色の油として得た。 A 250 mL round bottom flask was charged with a solution of 3-(benzyloxy)cyclobutan-1-one (6.0 g, 34.05 mmol, 1.0 equiv) in ethanol (100 mL). This was followed by the addition of sodium borohydride (1.25 g, 33.94 mmol, 1.00 equiv) in portions at 0° C. The resulting solution was stirred at 0° C. for 3 hours. The reaction was then quenched by the addition of water (100 mL). The resulting solution was extracted with ethyl acetate (50 mL×2) and the organic layers were combined, dried over anhydrous sodium sulfate, and concentrated under vacuum. This afforded 6.0 g (99%) of (1s,3s)-3-(benzyloxy)cyclobutan-1-ol as a pale yellow oil.
工程2

窒素不活性雰囲気下でパージし、維持した500mLの丸底フラスコに、N,N-ジメチルホルムアミド(100mL)中の(1s,3s)-3-(ベンジルオキシ)シクロブタン-1-オール(8.0g、44.89mmol、1.10当量)の溶液を入れた。続いて、水素化ナトリウム(60%)(2.15g、89.58mmol、1.20当量)を0℃で数回に分けて添加した。これに、4-ブロモ-2-フルオロピリジン(70.0g、39.78mmol、1.00当量)を添加した。得られた溶液を、室温で一晩撹拌した。次いで、反応物を、水/氷を添加することによってクエンチした。得られた溶液を酢酸エチル(80mL×2)で抽出し、有機層を合わせた。得られた混合物をブライン(80mL×2)で洗浄した。結果として生じる混合物を減圧下で濃縮した。残渣を、酢酸エチル/石油エーテル(1:1)で溶出するシリカゲルカラム上に適用した。これにより、7.0g(53%)の4-ブロモ-2-[(1s,3s)-3-(ベンジルオキシ)シクロブトキシ]ピリジンを淡い黄色の油として得た。 A 500 mL round bottom flask purged and maintained under nitrogen inert atmosphere was charged with a solution of (1s,3s)-3-(benzyloxy)cyclobutan-1-ol (8.0 g, 44.89 mmol, 1.10 equiv.) in N,N-dimethylformamide (100 mL). Sodium hydride (60%) (2.15 g, 89.58 mmol, 1.20 equiv.) was then added in portions at 0°C. To this was added 4-bromo-2-fluoropyridine (70.0 g, 39.78 mmol, 1.00 equiv.). The resulting solution was stirred overnight at room temperature. The reaction was then quenched by adding water/ice. The resulting solution was extracted with ethyl acetate (80 mL x 2) and the organic layers were combined. The resulting mixture was washed with brine (80 mL x 2). The resulting mixture was concentrated under reduced pressure. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (1:1). This gave 7.0 g (53%) of 4-bromo-2-[(1s,3s)-3-(benzyloxy)cyclobutoxy]pyridine as a pale yellow oil.
工程3
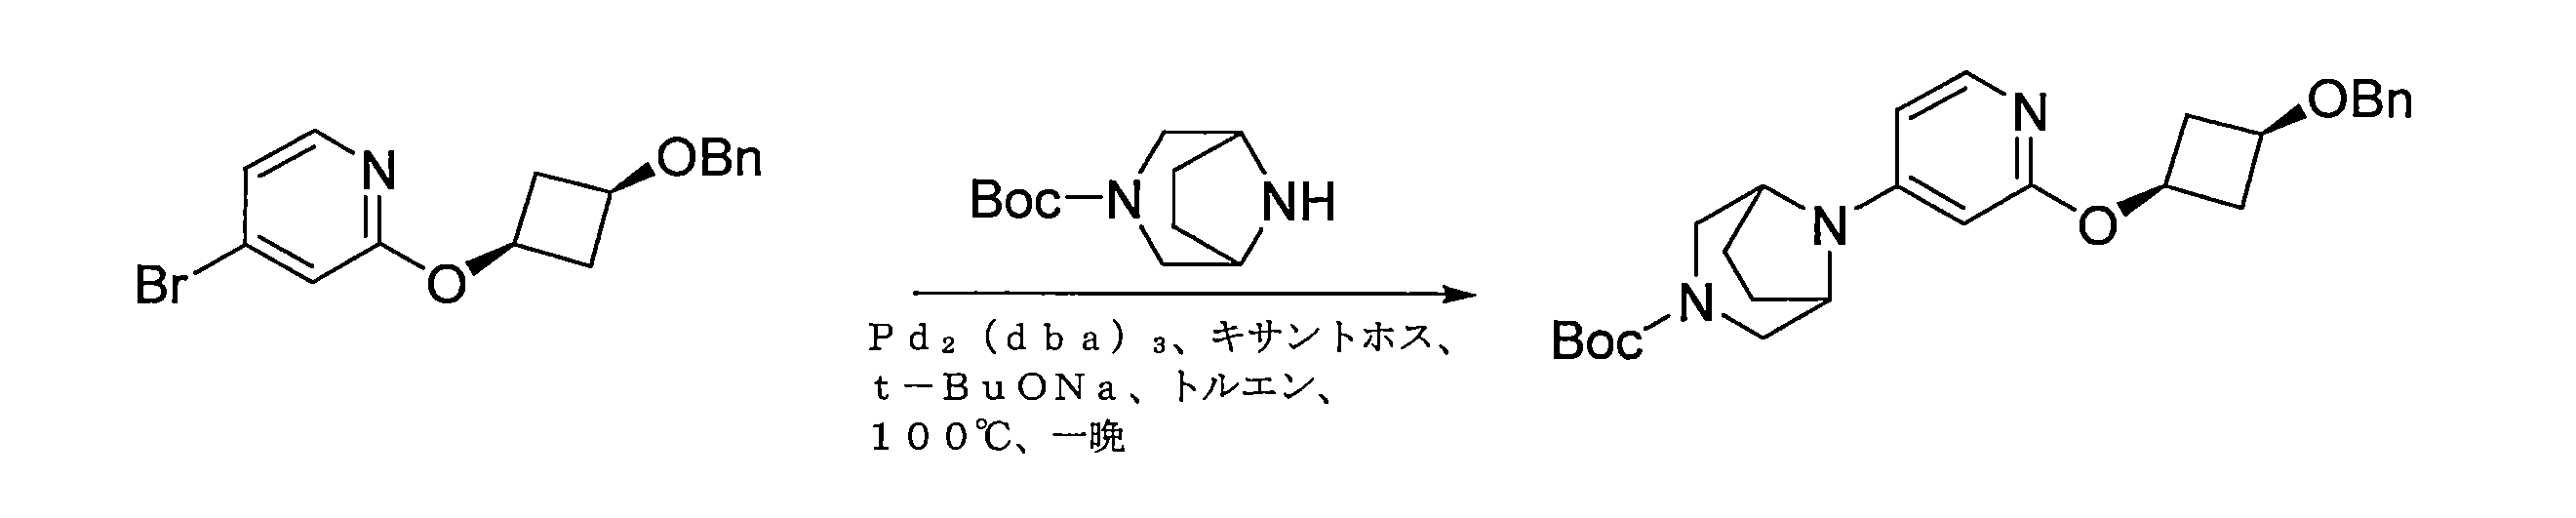
窒素不活性雰囲気下でパージし、維持した500mLの丸底フラスコに、トルエン(200mL)中の4-ブロモ-2-[(1s,3s)-3-(ベンジルオキシ)シクロブトキシ]ピリジン(8.4g、25.13mmol、1.30当量)の溶液、tert-ブチル3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレート(4.11g、19.36mmol、1.00当量)、tert-ブトキシドナトリウム(3.2g、33.2mmol、1.5当量)、キサントホス(2.32g、4.01mmol、0.20当量)、およびPd2(dba)3CH2Cl2(430.0mg、0.40mmol、0.02当量)を入れた。得られた溶液を100℃で一晩撹拌した。次いで、反応物を、水(100mL)を添加することによってクエンチした。得られた溶液を酢酸エチルで抽出し(80mL×2)、有機層を合わせた。得られた混合物をブライン(100mL×2)で洗浄した。混合物を無水硫酸ナトリウムで乾燥させ、減圧下で濃縮した。残渣を、酢酸エチル/ヘキサン(1:1)で溶出するシリカゲルカラム上に適用した。これにより、8.0g(89%)のtert-ブチル8-[2-[(1s,3s)-3-(ベンジルオキシ)シクロブトキシ]ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレートを淡い黄色の油として得た。 A 500 mL round bottom flask purged and maintained under a nitrogen inert atmosphere was charged with a solution of 4-bromo-2-[(1s,3s)-3-(benzyloxy)cyclobutoxy]pyridine (8.4 g, 25.13 mmol, 1.30 equiv.), tert-butyl 3,8-diazabicyclo[3.2.1]octane-3-carboxylate (4.11 g, 19.36 mmol, 1.00 equiv.), sodium tert-butoxide (3.2 g, 33.2 mmol, 1.5 equiv.), Xantphos (2.32 g, 4.01 mmol, 0.20 equiv.), and Pd2(dba)3CH2Cl2 (430.0 mg, 0.40 mmol, 0.02 equiv.) in toluene (200 mL). The resulting solution was stirred at 100 °C overnight. The reaction was then quenched by adding water (100 mL). The resulting solution was extracted with ethyl acetate (80 mL x 2) and the organic layers were combined. The resulting mixture was washed with brine (100 mL x 2). The mixture was dried over anhydrous sodium sulfate and concentrated under reduced pressure. The residue was applied onto a silica gel column eluted with ethyl acetate/hexane (1:1). This gave 8.0 g (89%) of tert-butyl 8-[2-[(1s,3s)-3-(benzyloxy)cyclobutoxy]pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octane-3-carboxylate as a pale yellow oil.
工程4

150mLのエタノール中のtert-ブチル8-[2-[(1s,3s)-3-(ベンジルオキシ)シクロブトキシ]ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレート(7.0g、15.03mmol、1.00当量)の溶液に、水酸化パラジウム(7.0g、49.85mmol、3.32当量)および酢酸(2mL)を、250mLの丸底フラスコ中で窒素雰囲気下で添加した。次いで、フラスコを真空化し、水素でフラッシュした。反応混合物を、水素バルーンを使用して水素雰囲気下で、50℃で3日間水素化し、次いで、セライトパッドを通して濾過し、減圧下で濃縮した。これにより、5.4g(96%)のtert-ブチル8-[2-[(1s,3s)-3-ヒドロキシシクロブトキシ]ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレートを淡い黄色の脂として得た。 To a solution of tert-butyl 8-[2-[(1s,3s)-3-(benzyloxy)cyclobutoxy]pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octane-3-carboxylate (7.0 g, 15.03 mmol, 1.00 equiv.) in 150 mL of ethanol, palladium hydroxide (7.0 g, 49.85 mmol, 3.32 equiv.) and acetic acid (2 mL) were added in a 250 mL round-bottom flask under nitrogen atmosphere. The flask was then evacuated and flushed with hydrogen. The reaction mixture was hydrogenated under hydrogen atmosphere using a hydrogen balloon at 50° C. for 3 days, then filtered through a celite pad and concentrated under reduced pressure. This yielded 5.4 g (96%) of tert-butyl 8-[2-[(1s,3s)-3-hydroxycyclobutoxy]pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octane-3-carboxylate as a pale yellow oil.
工程5

100mLの丸底フラスコに、酢酸エチルおよびメタノール(10:1)(50mL)の混合溶媒中のtert-ブチル8-[2-[(1s,3s)-3-ヒドロキシシクロブトキシ]ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレート(5.0g、13.32mmol、1.00当量)の溶液を入れた。塩化水素(ガス)を溶液に通した。得られた混合物を室温で3時間撹拌し、減圧下で濃縮した。これにより、4.0g(96%)の(1s,3s)-3-[(4-[3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)オキシ]シクロブタン-1-オール塩酸塩を灰色がかった白色の固体として得た。 A 100 mL round bottom flask was charged with a solution of tert-butyl 8-[2-[(1s,3s)-3-hydroxycyclobutoxy]pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octane-3-carboxylate (5.0 g, 13.32 mmol, 1.00 equiv.) in a mixture of ethyl acetate and methanol (10:1) (50 mL). Hydrogen chloride (gas) was passed through the solution. The resulting mixture was stirred at room temperature for 3 hours and concentrated under reduced pressure. This gave 4.0 g (96%) of (1s,3s)-3-[(4-[3,8-diazabicyclo[3.2.1]octane-8-yl]pyridin-2-yl)oxy]cyclobutan-1-ol hydrochloride as an off-white solid.
工程6

窒素の不活性雰囲気でパージし、維持した20mLの圧力タンク反応器に、ジメチルスルホキシド(10mL)中の(1s,3s)-3-[(4-[3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)オキシ]シクロブタン-1-オール塩酸塩(1.0g、3.21mmol、1.00当量)の溶液を入れた。次いで、N,N-ジイソプロピルエチルアミン(5mL)および4-ブロモ-6-クロロピリダジン-3-アミン(3.01g、14.44mmol、4.00当量)を添加した。得られた溶液を130℃で16時間撹拌した。次いで、反応物を、水(50mL)を添加することによってクエンチした。得られた溶液を酢酸エチル(3×30mL)で抽出し、有機層を合わせた。得られた混合物をブライン(3×50mL)で洗浄した。混合物を、無水硫酸ナトリウムで乾燥させた。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。これにより、490mg(38%)の(1s,3s)-3-([4-[3-(3-アミノ-6-クロロピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル]オキシ)シクロブタン-1-オールを黄色の油として得た。 A 20 mL pressure tank reactor purged and maintained with an inert atmosphere of nitrogen was charged with a solution of (1s,3s)-3-[(4-[3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl)oxy]cyclobutan-1-ol hydrochloride (1.0 g, 3.21 mmol, 1.00 equiv.) in dimethylsulfoxide (10 mL). N,N-diisopropylethylamine (5 mL) and 4-bromo-6-chloropyridazin-3-amine (3.01 g, 14.44 mmol, 4.00 equiv.) were then added. The resulting solution was stirred at 130° C. for 16 hours. The reaction was then quenched by adding water (50 mL). The resulting solution was extracted with ethyl acetate (3×30 mL) and the organic layers were combined. The resulting mixture was washed with brine (3×50 mL). The mixture was dried over anhydrous sodium sulfate. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (10:1). This gave 490 mg (38%) of (1s,3s)-3-([4-[3-(3-amino-6-chloropyridazin-4-yl)-3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl]oxy)cyclobutan-1-ol as a yellow oil.
工程7
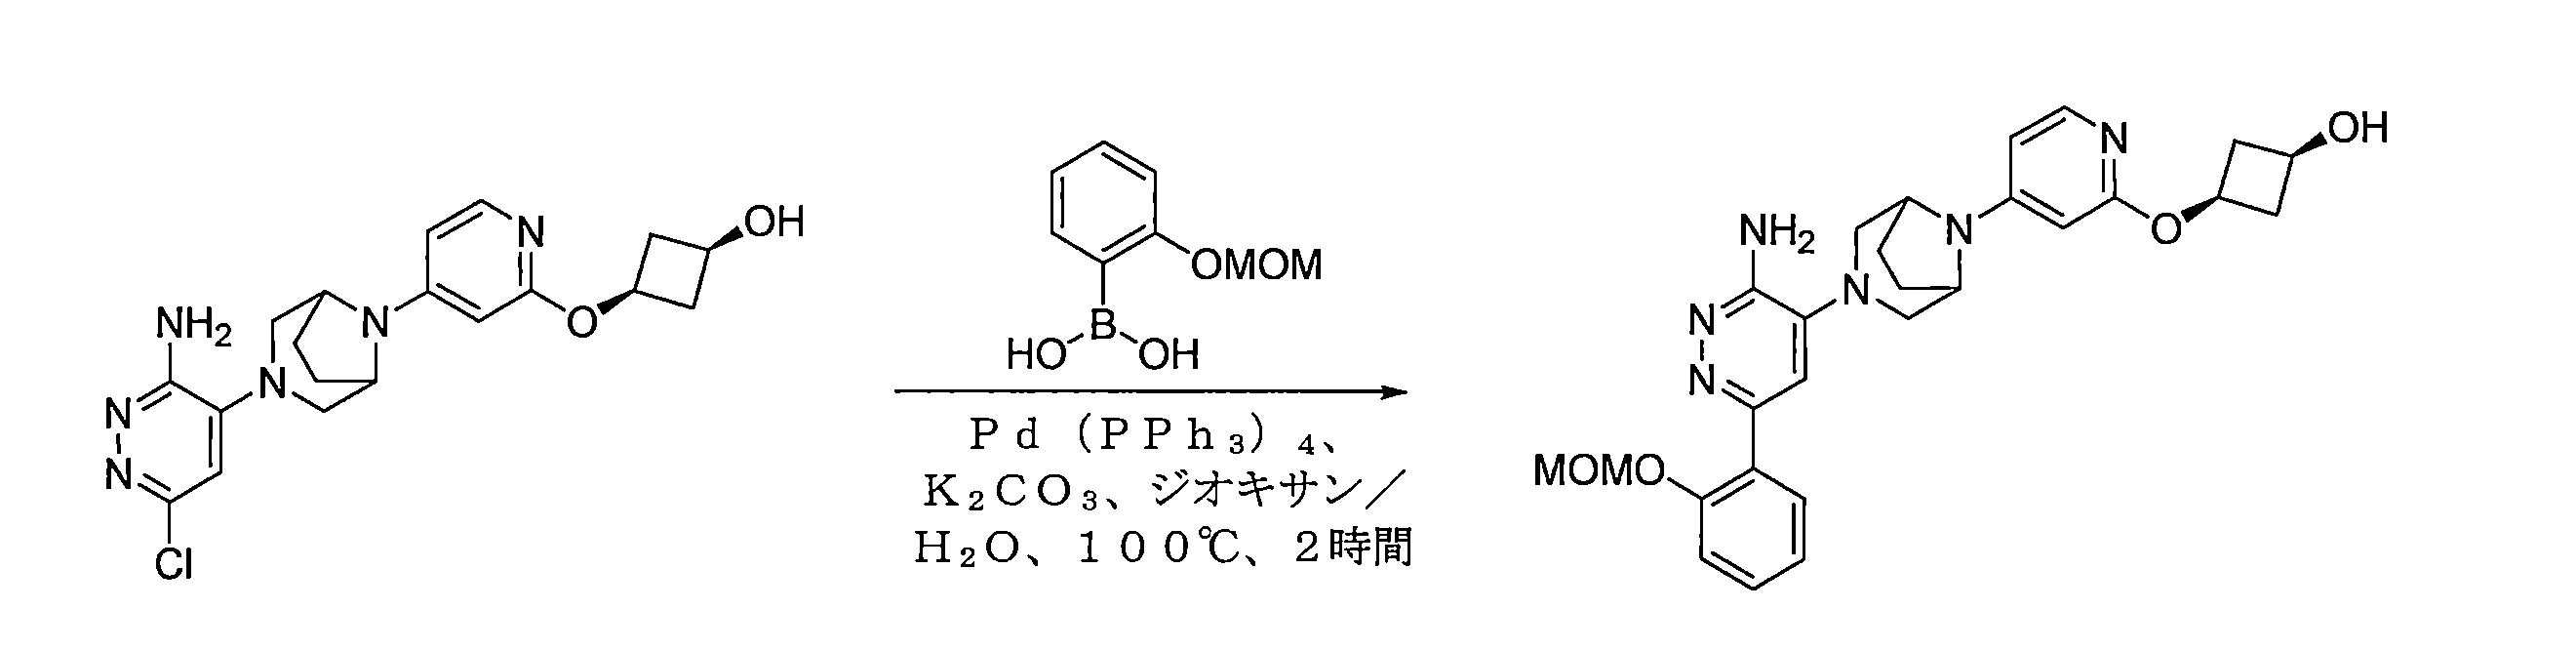
窒素不活性雰囲気下でパージし、維持した20mLの圧力タンク反応器に、ジオキサンと水(4:1)の混合溶媒(15mL)中の(1s,3s)-3-([4-[3-(3-アミノ-6-クロロピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル]オキシ)シクロブタン-1-オール(800mg、1.99mmol、1.00当量)の溶液、[2-(メトキシメトキシ)フェニル]ボロン酸(544mg、2.99mmol、1.50当量)、炭酸カリウム(824mg、6.0mmol、3.00当量)、およびPd(PPh3)4(230mg、0.20mmol、0.10当量)を入れた。得られた溶液を90℃で2時間撹拌した。次いで、反応物を、水(30mL)を添加することによってクエンチした。得られた溶液を酢酸エチル(50mL×2)で抽出し、有機層を合わせた。得られた混合物をブライン(50mL×2)で洗浄した。混合物を、無水硫酸ナトリウムで乾燥させた。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。これにより、820mg(82%)の(1s,3s)-3-[[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]オキシ]シクロブタン-1-オールを黄色の固体として得た。 A 20 mL pressure tank reactor purged and maintained under a nitrogen inert atmosphere was charged with a solution of (1s,3s)-3-([4-[3-(3-amino-6-chloropyridazin-4-yl)-3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl]oxy)cyclobutan-1-ol (800 mg, 1.99 mmol, 1.00 equiv.), [2-(methoxymethoxy)phenyl]boronic acid (544 mg, 2.99 mmol, 1.50 equiv.), potassium carbonate (824 mg, 6.0 mmol, 3.00 equiv.), and Pd(PPh3)4 (230 mg, 0.20 mmol, 0.10 equiv.) in a mixed solvent (15 mL) of dioxane and water (4:1). The resulting solution was stirred at 90° C. for 2 h. The reaction was then quenched by the addition of water (30 mL). The resulting solution was extracted with ethyl acetate (50 mL x 2) and the organic layers were combined. The resulting mixture was washed with brine (50 mL x 2). The mixture was dried over anhydrous sodium sulfate. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (10:1). This gave 820 mg (82%) of (1s,3s)-3-[[4-(3-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl]oxy]cyclobutan-1-ol as a yellow solid.
工程8
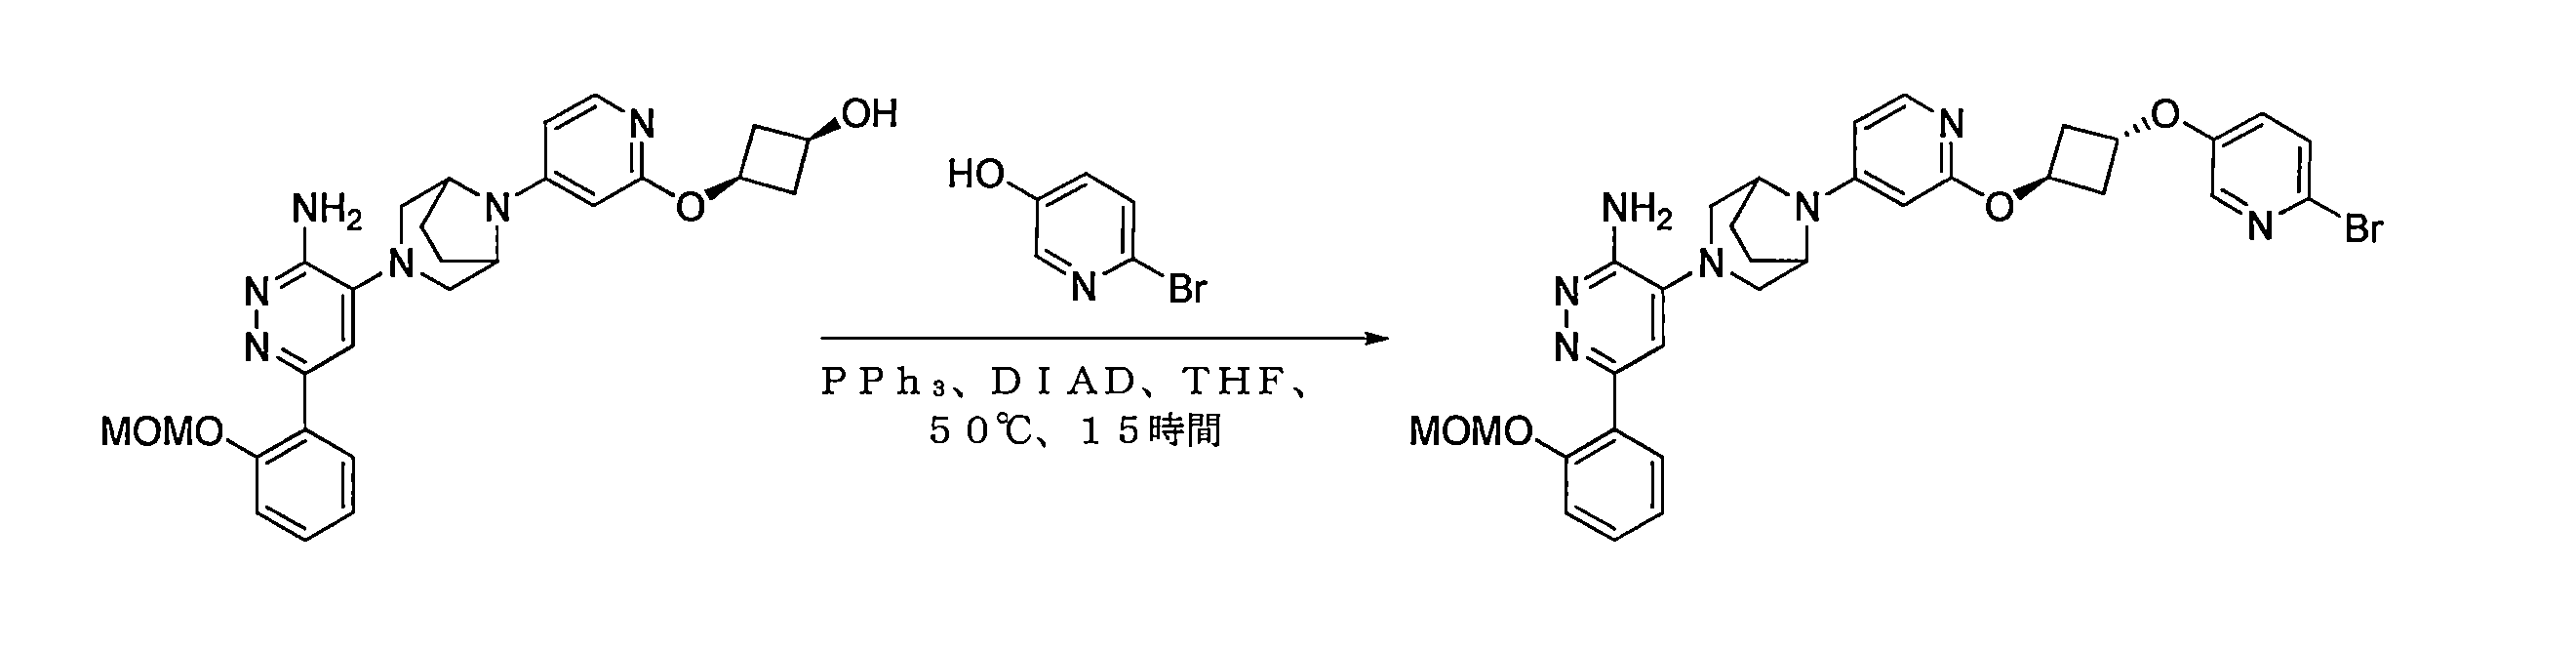
不活性雰囲気下でパージし、維持した50mLの三口丸底フラスコに、テトラヒドロフラン(蒸留)(30mL)中のトリフェニルホスフィン(351mg、1.34mmol、2.00当量)の溶液を入れた。これに続いて、ジイソプロピルアゾジカルボキシレート(270mg、1.34mmol、2.00当量)を、0℃で撹拌しながら滴下で添加し、白色の沈殿物が出現した。これに、(1s,3s)-3-[[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]オキシ]シクロブタン-1-オール(338mg、0.67mmol、1.00当量)および6-ブロモピリジン-3-オール(232mg、1.33mmol、1.50当量)を0℃で添加した。得られた溶液を50℃で一晩撹拌した。得られた混合物を真空下で濃縮した。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。これにより、389mg(88%)の6-[2-(メトキシメトキシ)フェニル]-4-(8-[2-[(1r,3r)-3-[(6-ブロモピリジン-3-イル)オキシ]シクロブトキシ]ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-イル)ピリダジン-3-アミンを黄色の固体として得た。 A 50 mL three-neck round bottom flask purged and maintained under an inert atmosphere was charged with a solution of triphenylphosphine (351 mg, 1.34 mmol, 2.00 equiv.) in tetrahydrofuran (distilled) (30 mL). This was followed by the dropwise addition of diisopropyl azodicarboxylate (270 mg, 1.34 mmol, 2.00 equiv.) with stirring at 0° C., resulting in the appearance of a white precipitate. To this was added (1s,3s)-3-[[4-(3-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl]oxy]cyclobutan-1-ol (338 mg, 0.67 mmol, 1.00 equiv.) and 6-bromopyridin-3-ol (232 mg, 1.33 mmol, 1.50 equiv.) at 0° C. The resulting solution was stirred at 50° C. overnight. The resulting mixture was concentrated under vacuum. The residue was applied onto a silica gel column eluting with dichloromethane/methanol (10:1). This gave 389 mg (88%) of 6-[2-(methoxymethoxy)phenyl]-4-(8-[2-[(1r,3r)-3-[(6-bromopyridin-3-yl)oxy]cyclobutoxy]pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octan-3-yl)pyridazin-3-amine as a yellow solid.
工程9
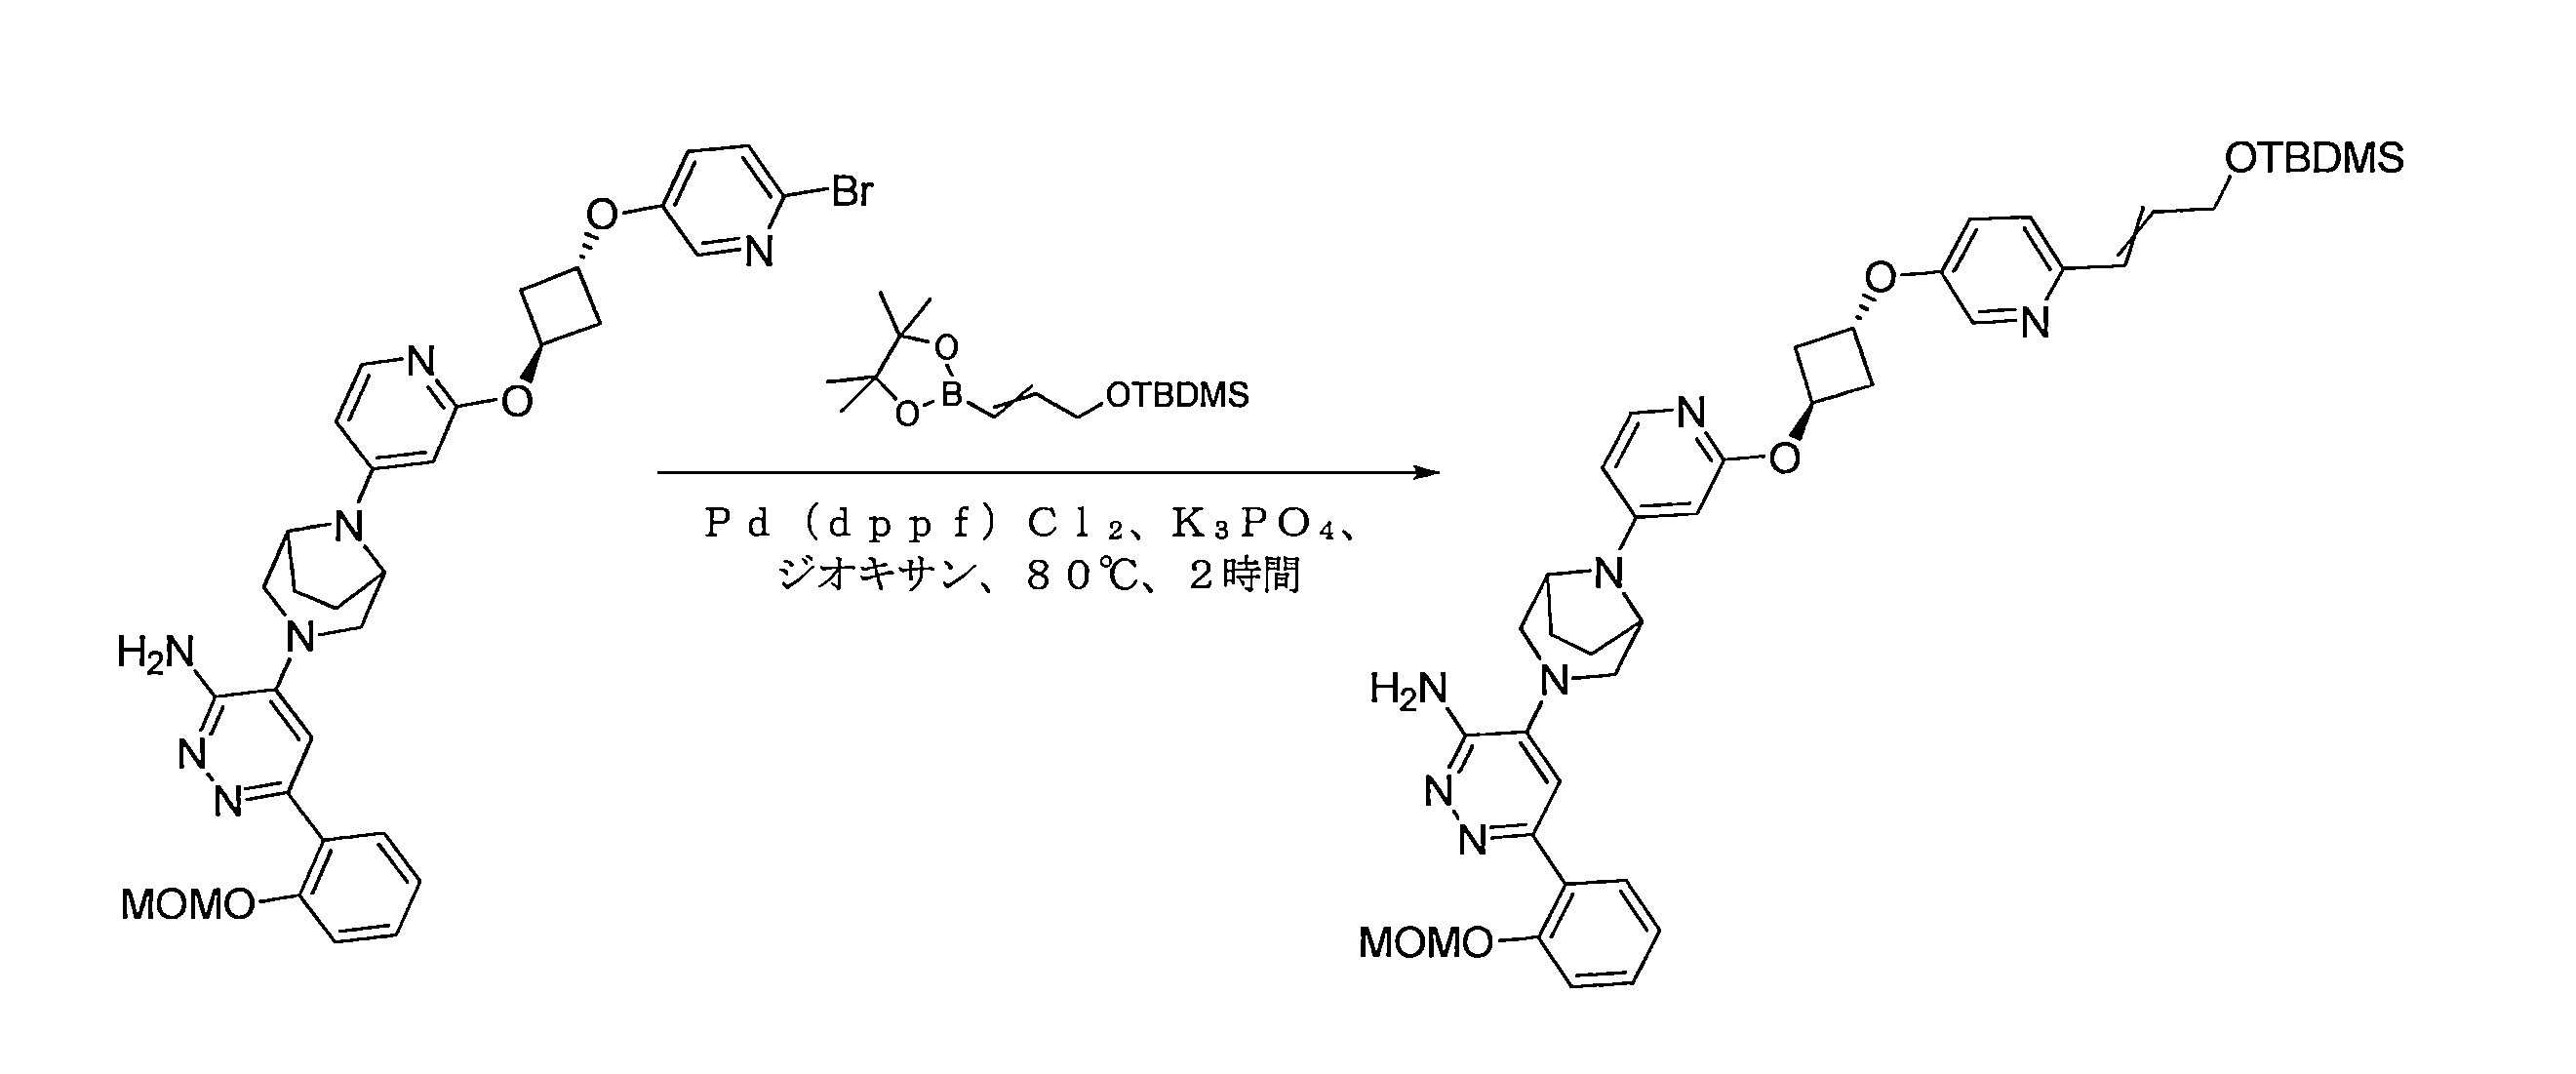
窒素不活性雰囲気下でパージし、維持した20mLの圧力タンク反応器に、ジオキサンと水(3:1)の混合溶媒(8mL)中の6-[2-(メトキシメトキシ)フェニル]-4-(8-[2-[(1r,3r)-3-[(6-ブロモピリジン-3-イル)オキシ]シクロブトキシ]ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-イル)ピリダジン-3-アミン(155mg、0.23mmol、1.00当量)の溶液、tert-ブチルジメチル{[3-(テトラメチル-1,3,2-ジオキサボロラン-2-イル)プロパ-2-エン-1-イル]オキシ}シラン(140mg、0.47mmol、2.00当量)、炭酸カリウム(95mg、0.69mmol、0.15当量)、Pd(PPh3)4Cl2CH2Cl2(28mg、0.03mmol、2.00当量)を入れた。得られた溶液を80℃で1時間撹拌した。次いで、反応物を、水を添加することによってクエンチした。得られた溶液を酢酸エチル(30mL×3)で抽出し、有機層を合わせた。得られた混合物をブライン(30mL×2)で洗浄した。混合物を、無水硫酸ナトリウムで乾燥させた。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。これにより、130mg(74%)の6-[2-(メトキシメトキシ)フェニル]-4-(8-[2-[(1r,3r)-3-[(6-[3-[(tert-ブチルジメチルシリル)オキシ]プロパ-1-エン-1-イル]ピリジン-3-イル)オキシ]シクロブトキシ]ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-イル)ピリダジン-3-アミンを淡い黄色の油として得た。 In a 20 mL pressure tank reactor purged and maintained under a nitrogen inert atmosphere, 155 mg of 6-[2-(methoxymethoxy)phenyl]-4-(8-[2-[(1r,3r)-3-[(6-bromopyridin-3-yl)oxy]cyclobutoxy]pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octan-3-yl)pyridazin-3-amine in a mixture of dioxane and water (3:1) (8 mL) was added. A solution of 3-(2,4-dioxaborolan-2-yl, 0.23 mmol, 1.00 equiv.), tert-butyldimethyl{[3-(tetramethyl-1,3,2-dioxaborolan-2-yl)prop-2-en-1-yl]oxy}silane (140 mg, 0.47 mmol, 2.00 equiv.), potassium carbonate (95 mg, 0.69 mmol, 0.15 equiv.), Pd(PPh3)4Cl2CH2Cl2 (28 mg, 0.03 mmol, 2.00 equiv.) was charged. The resulting solution was stirred at 80° C. for 1 h. The reaction was then quenched by adding water. The resulting solution was extracted with ethyl acetate (30 mL×3) and the organic layers were combined. The resulting mixture was washed with brine (30 mL×2). The mixture was dried over anhydrous sodium sulfate. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (10:1). This gave 130 mg (74%) of 6-[2-(methoxymethoxy)phenyl]-4-(8-[2-[(1r,3r)-3-[(6-[3-[(tert-butyldimethylsilyl)oxy]prop-1-en-1-yl]pyridin-3-yl)oxy]cyclobutoxy]pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octan-3-yl)pyridazin-3-amine as a pale yellow oil.
工程10

25mLの丸底フラスコに、テトラヒドロフラン(2mL)中の6-[2-(メトキシメトキシ)フェニル]-4-(8-[2-[(1r,3r)-3-[(6-[3-[(tert-ブチルジメチルシリル)オキシ]プロパ-1-エン-1-イル]ピリジン-3-イル)オキシ]シクロブトキシ]ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-イル)ピリダジン-3-アミン(120mg、0.16mmol、1.00当量)の溶液、フッ化テトラブチルアンモニウム(テトラヒドロフラン中1M)(3.2mL、2.00当量)を入れた。得られた溶液を室温で2時間撹拌した。次いで、反応を、水の添加により停止させた。得られた溶液を、酢酸エチル(30mL×3)で抽出し、有機層を合わせて、無水硫酸ナトリウムで乾燥させた。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。粗生成物を、以下の条件を用いて、フラッシュ分取HPLC(CombiFlash(登録商標)-1)によってさらに精製した。カラム、C18シリカゲル;移動相、水中のACN、30分以内に0%ACNが40%ACNに増加;検出器、UV220nm。これにより、70mg(69%)の3-[5-[(1r,3r)-3-[[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]オキシ]シクロブトキシ]ピリジン-2-イル]プロパ-2-エン-1-オールを淡い黄色の固体として得た。 A 25 mL round bottom flask was charged with a solution of 6-[2-(methoxymethoxy)phenyl]-4-(8-[2-[(1r,3r)-3-[(6-[3-[(tert-butyldimethylsilyl)oxy]prop-1-en-1-yl]pyridin-3-yl)oxy]cyclobutoxy]pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octan-3-yl)pyridazin-3-amine (120 mg, 0.16 mmol, 1.00 equiv.) in tetrahydrofuran (2 mL), tetrabutylammonium fluoride (1 M in tetrahydrofuran) (3.2 mL, 2.00 equiv.). The resulting solution was stirred at room temperature for 2 hours. The reaction was then quenched by the addition of water. The resulting solution was extracted with ethyl acetate (30 mL x 3) and the organic layers were combined and dried over anhydrous sodium sulfate. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (10:1). The crude product was further purified by flash preparative HPLC (CombiFlash®-1) using the following conditions: column, C18 silica gel; mobile phase, ACN in water, 0% ACN increasing to 40% ACN within 30 min; detector, UV 220 nm. This gave 70 mg (69%) of 3-[5-[(1r,3r)-3-[[4-(3-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl]oxy]cyclobutoxy]pyridin-2-yl]prop-2-en-1-ol as a pale yellow solid.
工程11

窒素不活性雰囲気下でパージし、維持した50mLの三口丸底フラスコに、テトラヒドロフラン(20mL)中のトリブチルホスフィン(149mg、0.74mmol、10.00当量)の溶液を入れた。これに続いて、テトラヒドロフラン(5mL)中のテトラメチルアゾジカルボキサミド(126mg、0.73mmol、10.00当量)の溶液を、-5℃で、撹拌しながら滴下で添加した。この混合物に、(2S,4R)-4-ヒドロキシ-1-[2-(3-ヒドロキシ-1,2-オキサゾール-5-イル)-3-メチルブタノイル]-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]ピロリジン-2-カルボキサミド(42mg、0.09mmol、1.20当量)、および(2E)-3-[5-[(1r,3r)-3-[[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]オキシ]シクロブトキシ]ピリジン-2-イル]プロパ-2-エン-1-オール(47mg、0.07mmol、1.00当量)を添加した。得られた溶液を50℃で一晩撹拌した。次いで、反応物を、水を添加することによってクエンチした。得られた溶液を酢酸エチル(30mL×3)で抽出し、有機層を合わせた。得られた混合物をブライン(50mL)で洗浄した。有機相を減圧下で蒸発させ、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。これにより、32mg(39%)の(2S,4R)-4-ヒドロキシ-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]-1-[3-メチル-2-(3-[[(2E)-3-[5-[(1r,3r)-3-[[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]オキシ]シクロブトキシ]ピリジン-2-イル]プロパ-2-エン-1-イル]オキシ]-1,2-オキサゾール-5-イル)ブタノイル]ピロリジン-2-カルボキサミドを淡い黄色の固体として得た。 A 50 mL three-neck round bottom flask purged and maintained under an inert atmosphere of nitrogen was charged with a solution of tributylphosphine (149 mg, 0.74 mmol, 10.00 equiv.) in tetrahydrofuran (20 mL). This was followed by the dropwise addition of a solution of tetramethylazodicarboxamide (126 mg, 0.73 mmol, 10.00 equiv.) in tetrahydrofuran (5 mL) at -5°C with stirring. To this mixture was added (2S,4R)-4-hydroxy-1-[2-(3-hydroxy-1,2-oxazol-5-yl)-3-methylbutanoyl]-N-[[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]pyrrolidine-2-carboxamide (42 mg, 0.09 mmol, 1.20 equiv.) and (2E)-3-[5-[(1r,3r)-3-[[4-(3-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl]oxy]cyclobutoxy]pyridin-2-yl]prop-2-en-1-ol (47 mg, 0.07 mmol, 1.00 equiv.). The resulting solution was stirred at 50° C. overnight. The reaction was then quenched by adding water. The resulting solution was extracted with ethyl acetate (30 mL x 3) and the organic layers were combined. The resulting mixture was washed with brine (50 mL). The organic phase was evaporated under reduced pressure and applied onto a silica gel column eluted with dichloromethane/methanol (10:1). This gave 32 mg (39%) of (2S,4R)-4-hydroxy-N-[[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]-1-[3-methyl-2-(3-[[(2E)-3-[5-[(1r,3r)-3-[[4-(3-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl]oxy]cyclobutoxy]pyridin-2-yl]prop-2-en-1-yl]oxy]-1,2-oxazol-5-yl)butanoyl]pyrrolidine-2-carboxamide as a pale yellow solid.
工程12

窒素不活性雰囲気下でパージし、維持した25mLの丸底フラスコに、メタノール(3mL)中の(2S,4R)-4-ヒドロキシ-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]-1-(3-メチル-2-[3-[(3-[5-[(1r,3r)-3-[[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]オキシ]シクロブトキシ]ピリジン-2-イル]プロパ-2-エン-1-イル)オキシ]-1,2-オキサゾール-5-イル]ブタノイル)ピロリジン-2-カルボキサミド(23mg、0.02mmol、1.00当量)の溶液および37%塩酸(1.5mL)を入れた。得られた溶液を室温で2時間撹拌した。反応混合物を、水/氷浴で0℃に冷却した。溶液のpH値を、NH4HCO3で7に調整した。得られた溶液をジクロロメタン(30mL×3)で抽出し、有機層を合わせた。得られた混合物をブライン(50mL×3)で洗浄した。混合物を、無水硫酸ナトリウムで乾燥させた。以下の条件の分取HPLCによって、粗生成物を精製した:カラム、XBridge Prep C18 OBDカラム、5um、19*150mm;移動相、水(10mmoL/LのNH4HCO3)およびACN(41.0%ACN、8分間で最大62.0%);検出器、UV254nm。これにより、8.1mg(37%)の(2S,4R)-4-ヒドロキシ-N-[[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]-1-(3-メチル-2-[3-[(3-[5-[(1r,3r)-3-[(4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)オキシ]シクロブトキシ]ピリジン-2-イル]プロパ-2-エン-1-イル)オキシ]-1,2-オキサゾール-5-イル]ブタノイル)ピロリジン-2-カルボキサミドを淡い黄色の固体として得た。 A 25 mL round bottom flask purged and maintained under an inert atmosphere of nitrogen was charged with a solution of (2S,4R)-4-hydroxy-N-[[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]-1-(3-methyl-2-[3-[(3-[5-[(1r,3r)-3-[[4-(3-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl]oxy]cyclobutoxy]pyridin-2-yl]prop-2-en-1-yl)oxy]-1,2-oxazol-5-yl]butanoyl)pyrrolidine-2-carboxamide (23 mg, 0.02 mmol, 1.00 equiv.) in methanol (3 mL) and 37% hydrochloric acid (1.5 mL). The resulting solution was stirred at room temperature for 2 hours. The reaction mixture was cooled to 0° C. with a water/ice bath. The pH value of the solution was adjusted to 7 with NH4HCO3. The resulting solution was extracted with dichloromethane (30 mL×3) and the organic layers were combined. The resulting mixture was washed with brine (50 mL×3). The mixture was dried over anhydrous sodium sulfate. The crude product was purified by preparative HPLC with the following conditions: column, XBridge Prep C18 OBD column, 5 um, 19*150 mm; mobile phase, water (10 mmol/L NH4HCO3) and ACN (41.0% ACN, max. 62.0% in 8 min); detector, UV 254 nm. This gave 8.1 mg (37%) of (2S,4R)-4-hydroxy-N-[[4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]-1-(3-methyl-2-[3-[(3-[5-[(1r,3r)-3-[(4-[3-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl)oxy]cyclobutoxy]pyridin-2-yl]prop-2-en-1-yl)oxy]-1,2-oxazol-5-yl]butanoyl)pyrrolidine-2-carboxamide as a pale yellow solid.
例示的な化合物57の例示的な合成
例示的な化合物57を、他の例示的な化合物の生成について上に記載される手順を使用して、以下のスキームに従って調製した。

例示的な化合物47の例示的な合成
工程1
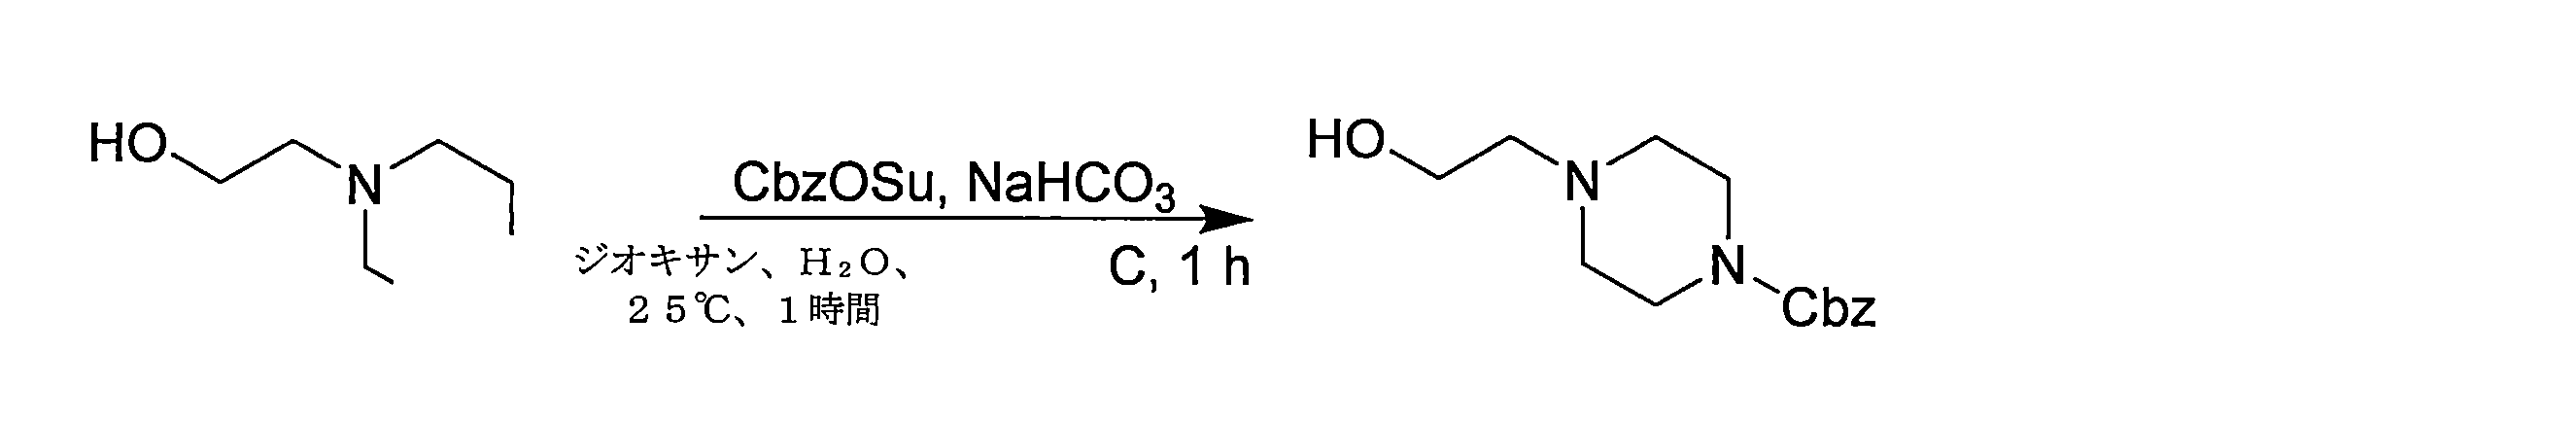
ジオキサン(300mL)および水(300mL)中の2-ピペラジン-1-イルエタノール(60g、460mmol、1当量)、重炭酸ナトリウム(116g、1.38mol、3当量)の溶液に、炭酸ベンジル(2,5-ジオキソピロリジン-1-イル)(137g、553mmol、1.2当量)を0℃で添加した。次いで、反応混合物を、20℃で2時間撹拌した。反応混合物を水(300mL)で希釈し、次いで、酢酸エチル(300mL×3)で抽出した。合わせた有機層をブライン(200mL×3)で洗浄し、無水硫酸ナトリウム上で乾燥させ、濾過し、減圧下で濃縮して残渣を得た。残渣を、カラムクロマトグラフィー(酢酸エチル:メタノール=500:1~100:1)によって精製した。化合物ベンジル4-(2-ヒドロキシエチル)ピペラジン-1-カルボキシレート(74g、279mmol、収率60%)を淡い黄色の油として得た。 To a solution of 2-piperazin-1-ylethanol (60 g, 460 mmol, 1 eq.), sodium bicarbonate (116 g, 1.38 mol, 3 eq.) in dioxane (300 mL) and water (300 mL), benzyl (2,5-dioxopyrrolidin-1-yl) carbonate (137 g, 553 mmol, 1.2 eq.) was added at 0°C. The reaction mixture was then stirred at 20°C for 2 hours. The reaction mixture was diluted with water (300 mL) and then extracted with ethyl acetate (300 mL x 3). The combined organic layers were washed with brine (200 mL x 3), dried over anhydrous sodium sulfate, filtered, and concentrated under reduced pressure to give a residue. The residue was purified by column chromatography (ethyl acetate:methanol = 500:1 to 100:1). The compound benzyl 4-(2-hydroxyethyl)piperazine-1-carboxylate (74 g, 279 mmol, 60% yield) was obtained as a pale yellow oil.
工程2
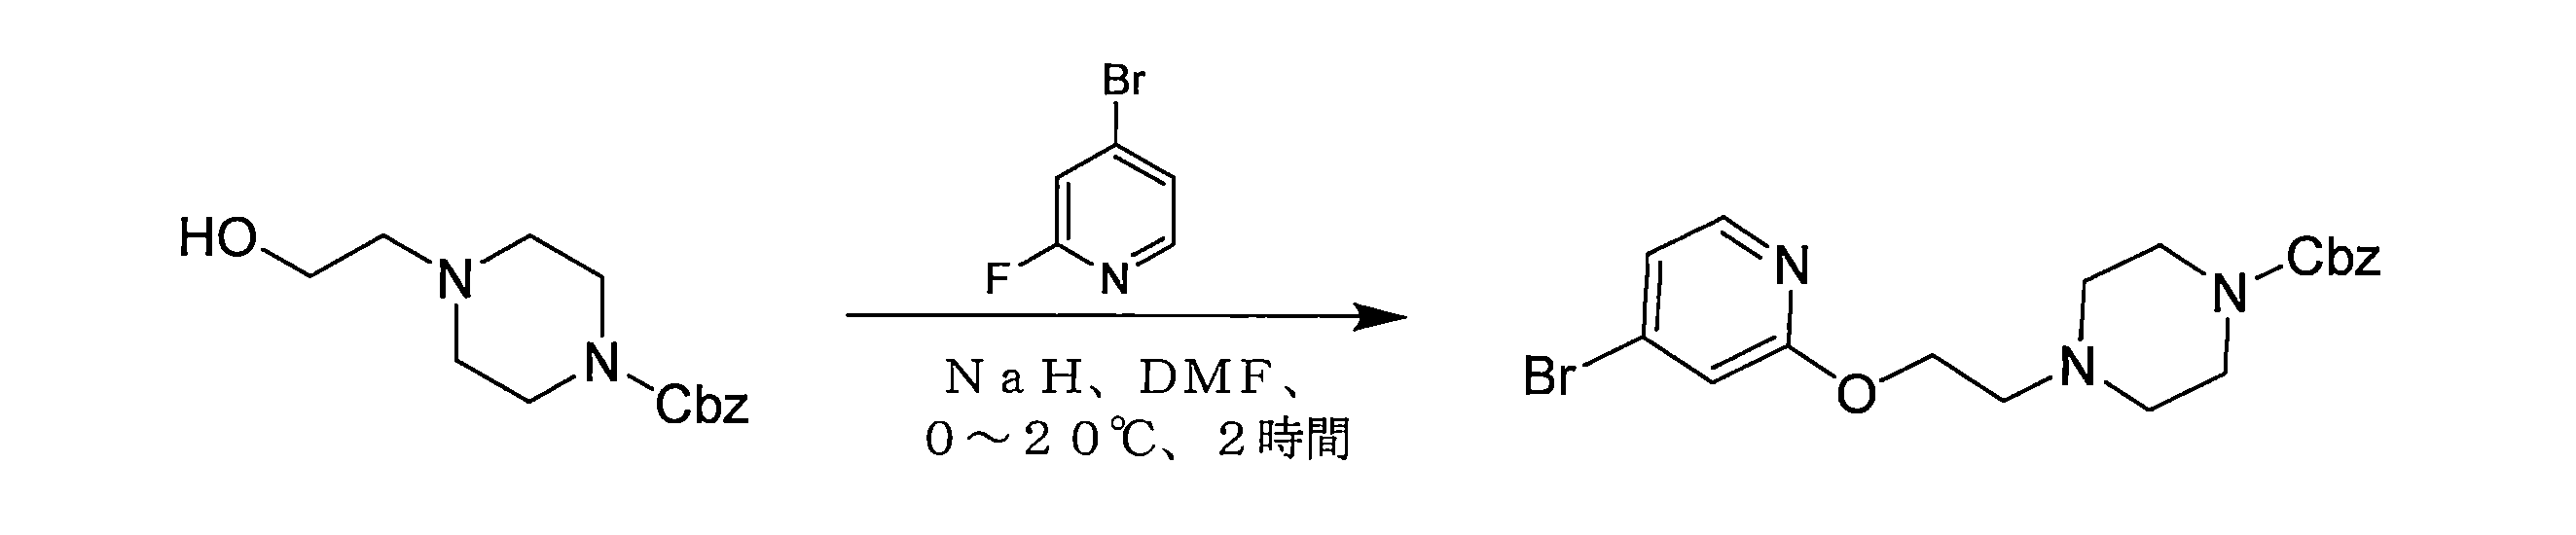
ジメチルホルムアミド(240mL)中の水素化ナトリウム(6.26g、156mmol、1.3当量)の溶液に、ベンジル4-(2-ヒドロキシエチル)ピペラジン-1-カルボキシレート(35g、132mmol、1.1当量)を0℃で0.5時間添加した。次いで、4-ブロモ-2-フルオロ-ピリジン(21.18g、120mmol、1当量)を反応物中に添加し、25℃で1時間撹拌した。反応混合物を、飽和塩化アンモニウム溶液(500mL)を添加することによってクエンチし、次いで、酢酸エチル(300mL×3)で抽出した。合わせた有機層をブライン(200mL×3)で洗浄し、無水硫酸ナトリウム上で乾燥させ、濾過し、減圧下で濃縮して残渣を得た。残渣を、カラムクロマトグラフィー(ジクロロメタン:メタノール=1:0~200:1)によって精製した。化合物ベンジル4-[2-[(4-ブロモ-2-ピリジル)オキシ]エチル]ピペラジン-1-カルボキシレート(39.2g、92mmol、収率76%)を淡い黄色の固体として得た。 To a solution of sodium hydride (6.26 g, 156 mmol, 1.3 equiv.) in dimethylformamide (240 mL), benzyl 4-(2-hydroxyethyl)piperazine-1-carboxylate (35 g, 132 mmol, 1.1 equiv.) was added at 0°C for 0.5 h. Then, 4-bromo-2-fluoro-pyridine (21.18 g, 120 mmol, 1 equiv.) was added into the reaction and stirred at 25°C for 1 h. The reaction mixture was quenched by adding saturated ammonium chloride solution (500 mL) and then extracted with ethyl acetate (300 mL x 3). The combined organic layers were washed with brine (200 mL x 3), dried over anhydrous sodium sulfate, filtered, and concentrated under reduced pressure to give a residue. The residue was purified by column chromatography (dichloromethane:methanol = 1:0 to 200:1). The compound benzyl 4-[2-[(4-bromo-2-pyridyl)oxy]ethyl]piperazine-1-carboxylate (39.2 g, 92 mmol, 76% yield) was obtained as a pale yellow solid.
工程3

バイアルに、ベンジル4-[2-[(4-ブロモ-2-ピリジル)オキシ]エチル]ピペラジン-1-カルボキシレート(18.15g、43mmol、1当量)、tert-ブチル3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレート(11g、51mmol、1.2当量)、[2-(2-アミノフェニル)フェニル]-クロロ-パラジウムジシクロヘキシル-[2-(2,6-ジイソプロポキシフェニル)フェニル]ホスファン(1.68g、2.16mmol、0.05当量)、炭酸セシウム(28.14g、86.36mmol、2当量)を充填し、トルエン(350mL)に添加した。混合物を、窒素で5分間パージし、次いで、110℃に12時間加熱した。反応混合物を水(200mL)で希釈し、酢酸エチル(300mL×3)で抽出した。合わせた有機層をブライン(200mL×3)で洗浄し、無水硫酸ナトリウム上で乾燥させ、濾過し、減圧下で濃縮して残渣を得た。残渣を、カラムクロマトグラフィー(石油エーテル/酢酸エチル=10:1~0:1)によって精製した。Tert-ブチル8-[2-[2-(4-ベンジルオキシカルボニルピペラジン-1-イル)エトキシ]-4-ピリジル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレート(18.37g、31.30mmol、収率72%)を淡い黄色の固体として得た。 A vial was charged with benzyl 4-[2-[(4-bromo-2-pyridyl)oxy]ethyl]piperazine-1-carboxylate (18.15 g, 43 mmol, 1 eq.), tert-butyl 3,8-diazabicyclo[3.2.1]octane-3-carboxylate (11 g, 51 mmol, 1.2 eq.), [2-(2-aminophenyl)phenyl]-chloro-palladium dicyclohexyl-[2-(2,6-diisopropoxyphenyl)phenyl]phosphane (1.68 g, 2.16 mmol, 0.05 eq.), cesium carbonate (28.14 g, 86.36 mmol, 2 eq.) and added to toluene (350 mL). The mixture was purged with nitrogen for 5 minutes and then heated to 110° C. for 12 hours. The reaction mixture was diluted with water (200 mL) and extracted with ethyl acetate (300 mL×3). The combined organic layers were washed with brine (200 mL x 3), dried over anhydrous sodium sulfate, filtered, and concentrated under reduced pressure to give a residue. The residue was purified by column chromatography (petroleum ether/ethyl acetate = 10:1 to 0:1). Tert-butyl 8-[2-[2-(4-benzyloxycarbonylpiperazin-1-yl)ethoxy]-4-pyridyl]-3,8-diazabicyclo[3.2.1]octane-3-carboxylate (18.37 g, 31.30 mmol, 72% yield) was obtained as a pale yellow solid.
工程4
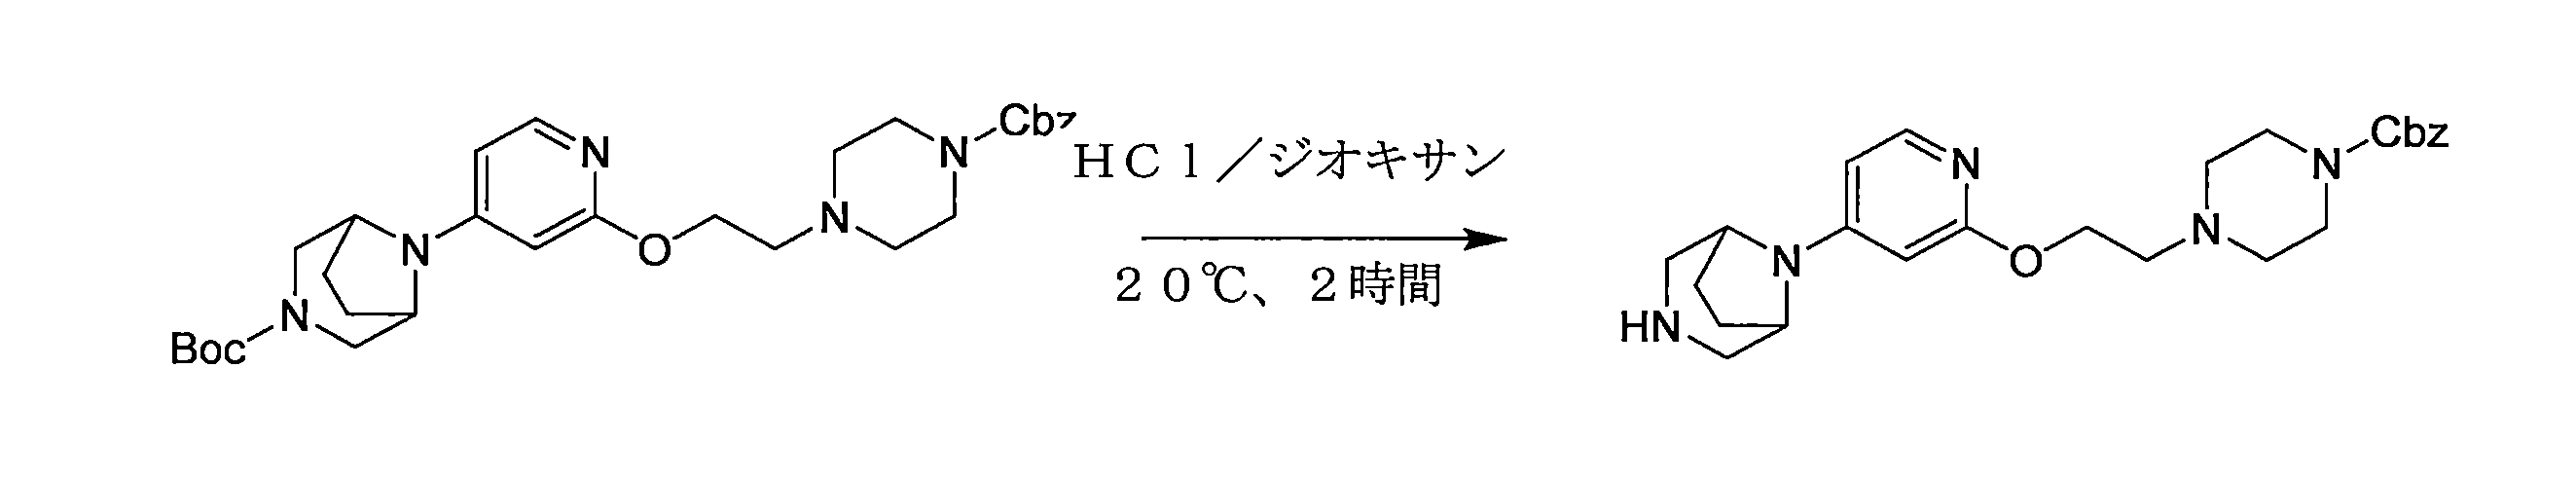
メタノール(150mL)中のtert-ブチル8-[2-[2-(4-ベンジルオキシカルボニルピペラジン-1-イル)エトキシ]-4-ピリジル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレート(18.3g、33.17mmol、1当量)の混合物に、塩酸/メタノール(4M、360mL、43.41当量)を添加した。次いで、反応混合物を、25℃で2時間撹拌した。反応混合物を減圧下で濃縮して、精製することなく化合物を得た。化合物ベンジル4-[2-[[4-(3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)-2-ピリジル]オキシ]エチル]ピペラジン-1-カルボキシレート(19g、粗製)を黄色の固体として得た。 To a mixture of tert-butyl 8-[2-[2-(4-benzyloxycarbonylpiperazin-1-yl)ethoxy]-4-pyridyl]-3,8-diazabicyclo[3.2.1]octane-3-carboxylate (18.3 g, 33.17 mmol, 1 eq.) in methanol (150 mL) was added hydrochloric acid/methanol (4 M, 360 mL, 43.41 eq.). The reaction mixture was then stirred at 25° C. for 2 hours. The reaction mixture was concentrated under reduced pressure to obtain the compound without purification. Compound benzyl 4-[2-[[4-(3,8-diazabicyclo[3.2.1]octane-8-yl)-2-pyridyl]oxy]ethyl]piperazine-1-carboxylate (19 g, crude) was obtained as a yellow solid.
工程5

ジメチルスルホキシド(190mL)中のベンジル4-[2-[[4-(3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)-2-ピリジル]オキシ]エチル]ピペラジン-1-カルボキシレート(19g、42.08mmol、1当量)、4-ブロモ-6-クロロ-ピリダジン-3-アミン(10.52g、50.49mmol、1.2当量)の混合物に、ジイソプロピルエチルアミン(54.38g、420.76mmol、10当量)を添加した。次いで、反応混合物を、130℃で5時間撹拌した。反応混合物を、水(600mL)を添加することによってクエンチし、次いで酢酸エチル(150mL)で希釈し、酢酸エチル(300mL×3)で抽出した。合わせた有機層をブライン(150mL×5)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、減圧下で濃縮して、残渣を得た。残渣を、カラムクロマトグラフィー(石油エーテル/酢酸エチル=1:1~ジクロロメタン:メタノール=50:1)によって精製した。化合物ベンジル4-[2-[[4-[3-(3-アミノ-6-クロロ-ピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]-2-ピリジル]オキシ]エチル]ピペラジン-1-カルボキシレート(12.6g、21mmol、収率50%)を淡い黄色の固体として得た。 To a mixture of benzyl 4-[2-[[4-(3,8-diazabicyclo[3.2.1]octan-8-yl)-2-pyridyl]oxy]ethyl]piperazine-1-carboxylate (19 g, 42.08 mmol, 1 eq.), 4-bromo-6-chloro-pyridazin-3-amine (10.52 g, 50.49 mmol, 1.2 eq.) in dimethyl sulfoxide (190 mL), diisopropylethylamine (54.38 g, 420.76 mmol, 10 eq.) was added. The reaction mixture was then stirred at 130° C. for 5 hours. The reaction mixture was quenched by adding water (600 mL), then diluted with ethyl acetate (150 mL) and extracted with ethyl acetate (300 mL×3). The combined organic layers were washed with brine (150 mL×5), dried over anhydrous sodium sulfate, filtered, and concentrated under reduced pressure to give a residue. The residue was purified by column chromatography (petroleum ether/ethyl acetate = 1:1 to dichloromethane:methanol = 50:1). The compound benzyl 4-[2-[[4-[3-(3-amino-6-chloro-pyridazin-4-yl)-3,8-diazabicyclo[3.2.1]octan-8-yl]-2-pyridyl]oxy]ethyl]piperazine-1-carboxylate (12.6 g, 21 mmol, 50% yield) was obtained as a pale yellow solid.
工程6

ジオキサン(160mL)および水(25mL)中のベンジル4-[2-[[4-[3-(3-アミノ-6-クロロ-ピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]-2-ピリジル]オキシ]エチル]ピペラジン-1-カルボキシレート(10.6g、18.30mmol、1当量)、(2-ヒドロキシフェニル)ボロン酸(5.05g、36.61mmol、2当量)の混合物に、テトラキス[トリフェニルホスフィン]パラジウム(0)(2.12g、1.83mmol、0.1当量)および炭酸カリウム(5.06g、36.61mmol、2当量)を添加した。次いで、反応混合物を、110℃で2時間撹拌した。反応混合物を、水(200mL)で希釈し、次いで、酢酸エチル(200mL×3)で抽出した。合わせた有機層をブライン(150mL×3)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、減圧下で濃縮して、残渣を得た。残渣を、分取HPLC(塩基条件:カラム:Phenomenex Gemini C18 250*50mm*10um;移動相:[水(0.05%水酸化アンモニアv/v)-アセトニトリル];B%:50%~80%、20分 60%分)によって精製し、所望の化合物を得た。化合物ベンジル4-[2-[[4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]-2-ピリジル]オキシ]エチル]ピペラジン-1-カルボキシレート(7g、10.22mmol、収率55%)を黄色の固体として得た。 To a mixture of benzyl 4-[2-[[4-[3-(3-amino-6-chloro-pyridazin-4-yl)-3,8-diazabicyclo[3.2.1]octan-8-yl]-2-pyridyl]oxy]ethyl]piperazine-1-carboxylate (10.6 g, 18.30 mmol, 1 eq.), (2-hydroxyphenyl)boronic acid (5.05 g, 36.61 mmol, 2 eq.) in dioxane (160 mL) and water (25 mL), tetrakis[triphenylphosphine]palladium(0) (2.12 g, 1.83 mmol, 0.1 eq.) and potassium carbonate (5.06 g, 36.61 mmol, 2 eq.) were added. The reaction mixture was then stirred at 110°C for 2 hours. The reaction mixture was diluted with water (200 mL) and then extracted with ethyl acetate (200 mL x 3). The combined organic layers were washed with brine (150 mL x 3), dried over anhydrous sodium sulfate, filtered and concentrated under reduced pressure to give a residue. The residue was purified by preparative HPLC (basic conditions: column: Phenomenex Gemini C18 250*50 mm*10 um; mobile phase: [water (0.05% ammonia hydroxide v/v)-acetonitrile]; B%: 50%-80%, 20 min 60% min) to give the desired compound. Compound benzyl 4-[2-[[4-[3-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl]-2-pyridyl]oxy]ethyl]piperazine-1-carboxylate (7 g, 10.22 mmol, 55% yield) was obtained as a yellow solid.
工程7

テトラヒドロフラン(100mL)中のベンジル4-[2-[[4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]-2-ピリジル]オキシ]エチル]ピペラジン-1-カルボキシレート(7g、10.99mmol、1当量)の溶液に、活性化炭素触媒上の水酸化パラジウム(0.7g、0.49mmol、0.04当量)を窒素下で添加した。次いで、反応混合物を水素(50Psi)下で、60℃において48時間撹拌した。次いで、150mLのエチルアルコールを反応混合物に添加した。反応混合物を濾過し、減圧下で濃縮して、2-[6-アミノ-5-[8-[2-(2-ピペラジン-1-イルエトキシ)-4-ピリジル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-イル]ピリダジン-3-イル]フェノール(5g、9.68mmol、収率88%)を得た。 To a solution of benzyl 4-[2-[[4-[3-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl]-2-pyridyl]oxy]ethyl]piperazine-1-carboxylate (7 g, 10.99 mmol, 1 equiv.) in tetrahydrofuran (100 mL) was added palladium hydroxide on activated carbon catalyst (0.7 g, 0.49 mmol, 0.04 equiv.) under nitrogen. The reaction mixture was then stirred under hydrogen (50 Psi) at 60° C. for 48 hours. 150 mL of ethyl alcohol was then added to the reaction mixture. The reaction mixture was filtered and concentrated under reduced pressure to give 2-[6-amino-5-[8-[2-(2-piperazin-1-ylethoxy)-4-pyridyl]-3,8-diazabicyclo[3.2.1]octan-3-yl]pyridazin-3-yl]phenol (5 g, 9.68 mmol, 88% yield).
工程8
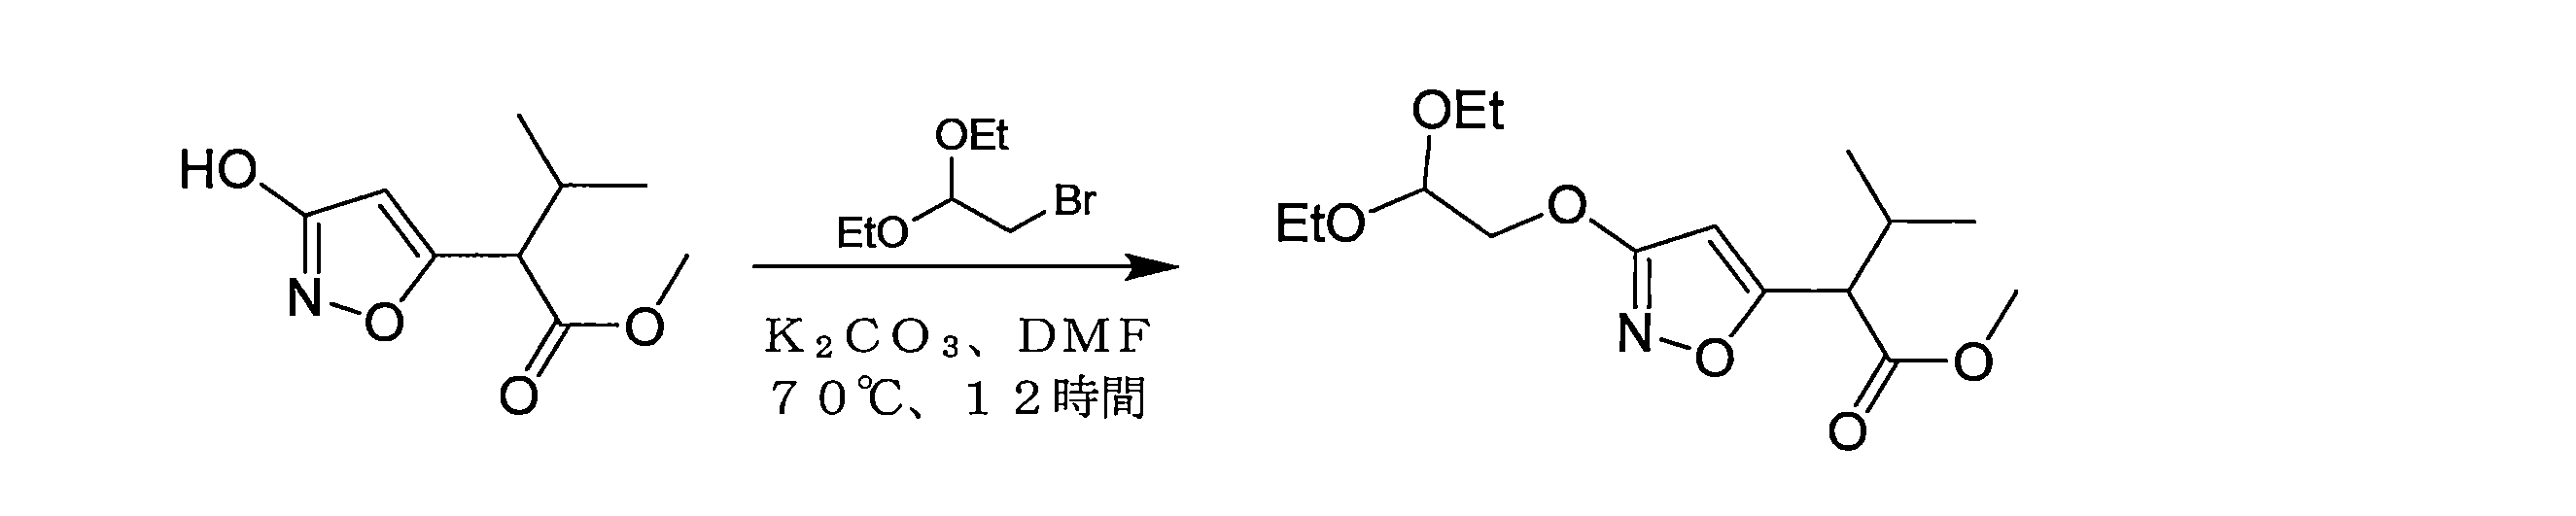
N,N-ジメチルホルムアミド(40mL)中のメチル2-(3-ヒドロキシイソキサゾール-5-イル)-3-メチル-ブタノエート(3.5g、17.57mmol、1当量)の溶液に、2-ブロモ-1,1-ジエトキシ-エタン(5.19g、26.35mmol、1.5当量)および炭酸カリウム(4.86g、35.14mmol、2当量)を添加した。反応混合物を70℃で12時間撹拌した。反応混合物を、水50mLを添加することによってクエンチし、次いで水100mLで希釈し、酢酸エチル(80mL×3)で抽出した。合わせた有機層を生理食塩水(100mL×2)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、減圧下で濃縮して、残渣を得た。残渣を、シリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=100:1~50:1)によって精製した。所望の化合物メチル2-[3-(2,2-ジエトキシエトキシ)イソキサゾール-5-イル]-3-メチル-ブタノエート(4.38g、12.50mmol、収率71%)を無色の油として得た。 To a solution of methyl 2-(3-hydroxyisoxazol-5-yl)-3-methyl-butanoate (3.5 g, 17.57 mmol, 1 eq.) in N,N-dimethylformamide (40 mL), 2-bromo-1,1-diethoxy-ethane (5.19 g, 26.35 mmol, 1.5 eq.) and potassium carbonate (4.86 g, 35.14 mmol, 2 eq.) were added. The reaction mixture was stirred at 70 °C for 12 h. The reaction mixture was quenched by adding 50 mL of water, then diluted with 100 mL of water and extracted with ethyl acetate (80 mL x 3). The combined organic layers were washed with saline (100 mL x 2), dried over anhydrous sodium sulfate, filtered, and concentrated under reduced pressure to give a residue. The residue was purified by silica gel chromatography (petroleum ether:ethyl acetate = 100:1 to 50:1). The desired compound, methyl 2-[3-(2,2-diethoxyethoxy)isoxazol-5-yl]-3-methyl-butanoate (4.38 g, 12.50 mmol, 71% yield), was obtained as a colorless oil.
工程9

メタノール(30mL)および水(15mL)中のメチル2-[3-(2,2-ジエトキシエトキシ)イソキサゾール-5-イル]-3-メチル-ブタノエート(4.38g、12.5mmol、1当量)の溶液に、水酸化リチウム一水和物(2.10g、50mmol、4当量)を添加した。反応混合物を、40℃で2時間撹拌した。pHを、1M塩化水素で4~5に調整し、反応混合物を酢酸エチル(50mL×2)で抽出した。合わせた有機層を無水硫酸ナトリウムで乾燥させ、濾過し、減圧下で濃縮して、2-[3-(2,2-ジエトキシエトキシ)イソキサゾール-5-イル]-3-メチル-ブタン酸(4g、粗製)を無色の油として得た。 To a solution of methyl 2-[3-(2,2-diethoxyethoxy)isoxazol-5-yl]-3-methyl-butanoate (4.38 g, 12.5 mmol, 1 equiv.) in methanol (30 mL) and water (15 mL) was added lithium hydroxide monohydrate (2.10 g, 50 mmol, 4 equiv.). The reaction mixture was stirred at 40°C for 2 h. The pH was adjusted to 4-5 with 1 M hydrogen chloride and the reaction mixture was extracted with ethyl acetate (50 mL x 2). The combined organic layers were dried over anhydrous sodium sulfate, filtered and concentrated under reduced pressure to give 2-[3-(2,2-diethoxyethoxy)isoxazol-5-yl]-3-methyl-butanoic acid (4 g, crude) as a colorless oil.
工程10

N,N-ジメチルホルムアミド(20mL)中の2-[3-(2,2-ジエトキシエトキシ)イソキサゾール-5-イル]-3-メチル-ブタン酸(3.84g、11.99mmol、1当量)の溶液に、O-(7-アザベンゾトリアゾール-1-イル)-N,N,N’,N’-テトラメチルウロニウムヘキサフルオロホスフェート(5.47g、14.38mmol、1.2当量)を添加した。反応混合物を、20℃で0.5時間撹拌した。次いで、混合物に、N,N-ジメチルホルムアミド(10mL)中の(2S,4R)-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド塩酸塩(4.41g、11.99mmol、1当量)およびトリエチルアミン(3.64g、35.96mmol、5mL、3当量)の溶液を添加した。反応混合物を、20℃で0.15時間撹拌した。反応混合物を、水(30mL)を添加することによってクエンチし、次いで水(80mL)で希釈し、酢酸エチル(60mL×3)で抽出した。合わせた有機層をブライン(80mL×2)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、減圧下で濃縮して、残渣を得た。残渣を、シリカゲルクロマトグラフィー(ジクロロメタン:メタノール=100:1~50:1)によって精製した。所望の化合物(2S,4R)-1-[2-[3-(2,2-ジエトキシエトキシ)イソキサゾール-5-イル]-3-メチル-ブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(7g、10.96mmol、収率91%)を白色の固体として得た。 To a solution of 2-[3-(2,2-diethoxyethoxy)isoxazol-5-yl]-3-methyl-butanoic acid (3.84 g, 11.99 mmol, 1 equiv.) in N,N-dimethylformamide (20 mL) was added O-(7-azabenzotriazol-1-yl)-N,N,N',N'-tetramethyluronium hexafluorophosphate (5.47 g, 14.38 mmol, 1.2 equiv.). The reaction mixture was stirred at 20°C for 0.5 h. To the mixture was then added a solution of (2S,4R)-4-hydroxy-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide hydrochloride (4.41 g, 11.99 mmol, 1 eq.) and triethylamine (3.64 g, 35.96 mmol, 5 mL, 3 eq.) in N,N-dimethylformamide (10 mL). The reaction mixture was stirred at 20° C. for 0.15 h. The reaction mixture was quenched by adding water (30 mL), then diluted with water (80 mL) and extracted with ethyl acetate (60 mL×3). The combined organic layers were washed with brine (80 mL×2), dried over anhydrous sodium sulfate, filtered and concentrated under reduced pressure to give a residue. The residue was purified by silica gel chromatography (dichloromethane:methanol=100:1 to 50:1). The desired compound (2S,4R)-1-[2-[3-(2,2-diethoxyethoxy)isoxazol-5-yl]-3-methyl-butanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (7 g, 10.96 mmol, 91% yield) was obtained as a white solid.
工程11

(2S,4R)-1-[2-[3-(2,2-ジエトキシエトキシ)イソキサゾール-5-イル]-3-メチル-ブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(7g、11.39mmol、1当量)をSFCによって分離した。条件は、カラム:DAICEL CHIRALPAK AD(250mm*30mm、10um);移動相:[0.1%NH3H2O IPA];B%:35%~35%、2.4分:550分。(2S,4R)-1-[(2S)-2-[3-(2,2-ジエトキシエトキシ)イソキサゾール-5-イル]-3-メチル-ブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(3.2g、5.08mmol、収率89%)および(2S,4R)-1-[(2R)-2-[3-(2,2-ジエトキシエトキシ)イソキサゾール-5-イル]-3-メチ
ル-ブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(2.67g、4.17mmol、収率73%)を白色の固体として得た。
(2S,4R)-1-[2-[3-(2,2-diethoxyethoxy)isoxazol-5-yl]-3-methyl-butanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (7 g, 11.39 mmol, 1 eq.) was separated by SFC. Conditions were: Column: DAICEL CHIRALPAK AD (250 mm*30 mm, 10 um); Mobile phase: [0.1% NH3H2O IPA]; B%: 35%-35%, 2.4 min: 550 min. (2S,4R)-1-[(2S)-2-[3-(2,2-diethoxyethoxy)isoxazol-5-yl]-3-methyl-butanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (3.2 g, 5.08 mmol, 89% yield) and (2S,4R)-1-[(2R)-2-[3-(2,2-diethoxyethoxy)isoxazol-5-yl]-3-methyl-butanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (2.67 g, 4.17 mmol, 73% yield) were obtained as white solids.
工程12

(2S,4R)-1-[(2R)-2-[3-(2,2-ジエトキシエトキシ)イソキサゾール-5-イル]-3-メチル-ブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(2g、3.25mmol、1当量)のテトラヒドロフラン(40mL)溶液に、を硫酸(1M、40mL、12.3当量)を添加した。溶液を、50℃に7時間加熱した。溶液を20℃に冷却し、飽和重炭酸ナトリウム溶液でpH=8までクエンチした。混合物を酢酸エチル(30mL×3)で抽出した。合わせた有機層をブライン(50mL)で洗浄し、硫酸ナトリウムで乾燥させた。次いで、混合物を濾過し、濾液を真空中で濃縮した。粗生成物を、次の工程で直接使用した。(2S,4R)-4-ヒドロキシ-1-[(2R)-3-メチル-2-[3-(2-オキソエトキシ)イソキサゾール-5-イル]ブタノイル]-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(1.7g、2.52mmol、収率77%)を白色の固体として得た。 To a solution of (2S,4R)-1-[(2R)-2-[3-(2,2-diethoxyethoxy)isoxazol-5-yl]-3-methyl-butanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (2 g, 3.25 mmol, 1 eq.) in tetrahydrofuran (40 mL) was added sulfuric acid (1 M, 40 mL, 12.3 eq.). The solution was heated to 50°C for 7 h. The solution was cooled to 20°C and quenched with saturated sodium bicarbonate solution to pH = 8. The mixture was extracted with ethyl acetate (30 mL x 3). The combined organic layers were washed with brine (50 mL) and dried over sodium sulfate. The mixture was then filtered and the filtrate was concentrated in vacuo. The crude product was used directly in the next step. (2S,4R)-4-hydroxy-1-[(2R)-3-methyl-2-[3-(2-oxoethoxy)isoxazol-5-yl]butanoyl]-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (1.7 g, 2.52 mmol, 77% yield) was obtained as a white solid.
工程13
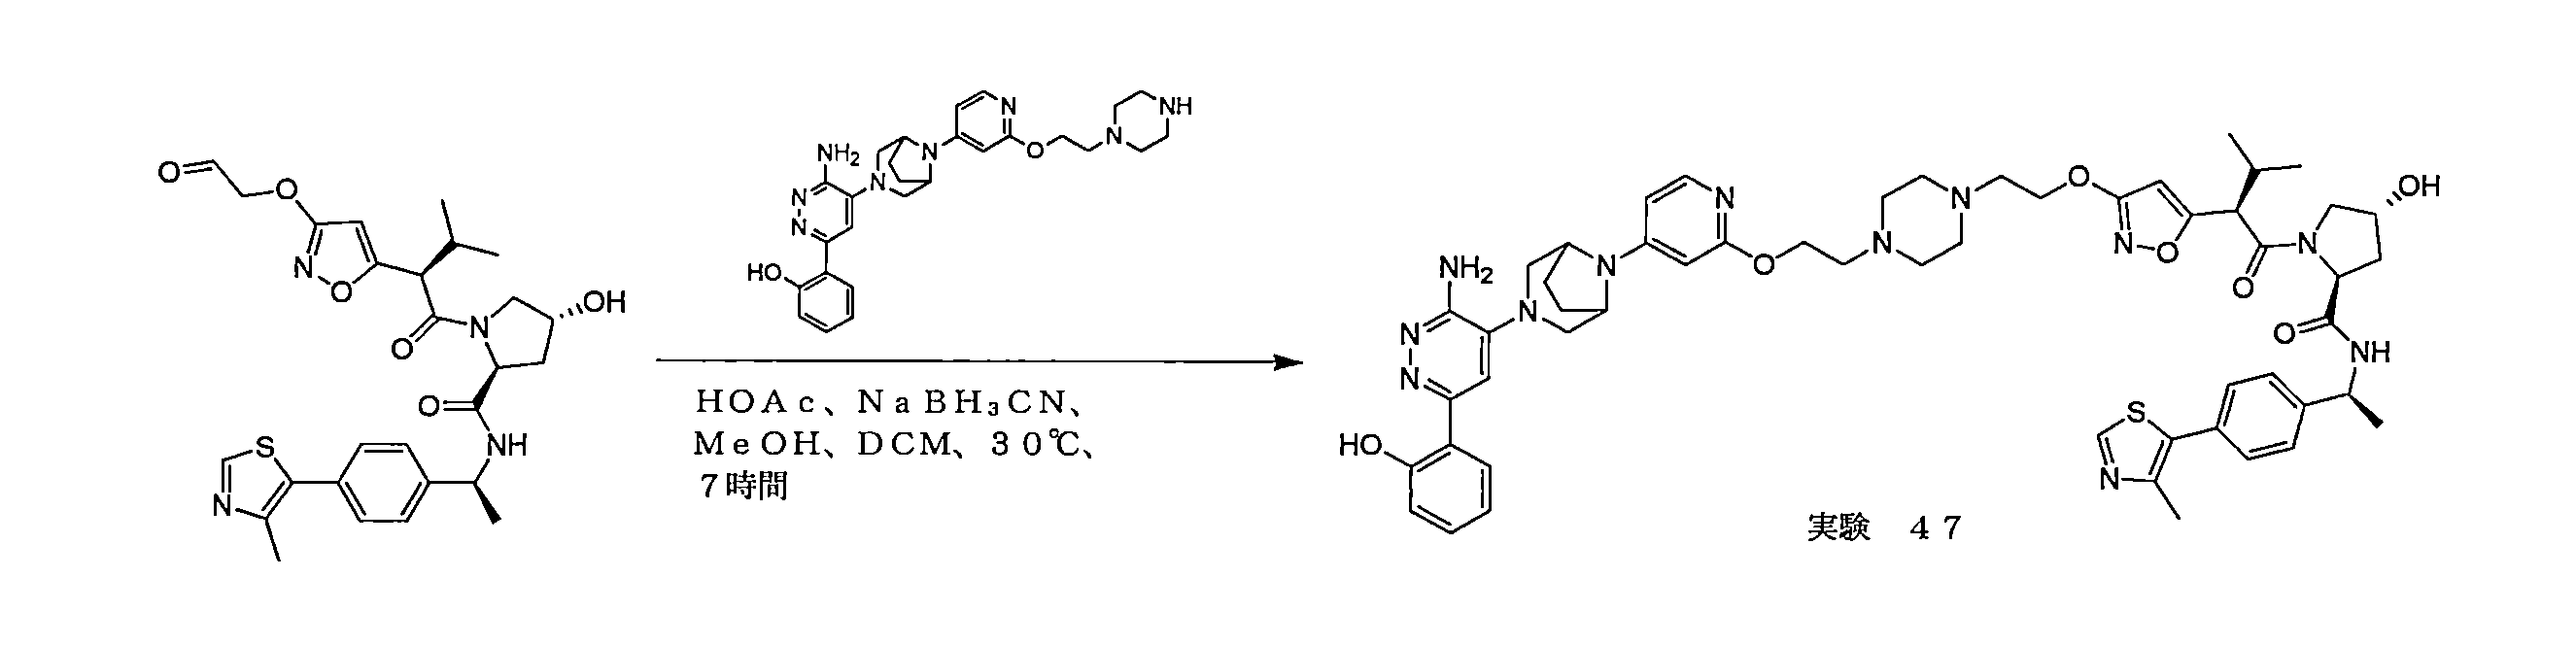
メタノール(15mL)およびジクロロメタン(5mL)中の(2S,4R)-4-ヒドロキシ-1-[(2R)-3-メチル-2-[3-(2-オキソエトキシ)イソキサゾール-5-イル]ブタノイル]-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(930mg、1.38mmol、1当量)および2-[6-アミノ-5-[8-[2-(2-ピペラジン-1-イルエトキシ)-4-ピリジル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-イル]ピリダジン-3-イル]フェノール(691mg、1.38mmol、1当量)の溶液に、酢酸(82mg、1.38mmol、1当量)を添加した。溶液を、30℃で0.5時間撹拌した。次いで、シアノ水素化ホウ素ナトリウム(129mg、2.06mmol、1.5当量)を溶液に添加し、30℃で7時間撹拌した。溶媒を、真空中で除去し、粗生成物を、分取HPLC(カラム:Phenomenex Synergi Max-RP 250*50mm*10um;移動相:[水(10mM NH4HCO3)-ACN];B%:45ACN%~75ACN%、28分、78%分)によって精製した。(2S,4R)-1-[(2R)-2-[3-[2-[4-[2-[[4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]-2-ピリジル]オキシ]エチル]ピペラジン-1-イル]エトキシ]イソキサゾール-5-イル]-3-メチル-ブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(840mg、0.8mmol、収率58%、純度98%)を白色の固体として得た。 To a solution of (2S,4R)-4-hydroxy-1-[(2R)-3-methyl-2-[3-(2-oxoethoxy)isoxazol-5-yl]butanoyl]-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (930 mg, 1.38 mmol, 1 equiv.) and 2-[6-amino-5-[8-[2-(2-piperazin-1-ylethoxy)-4-pyridyl]-3,8-diazabicyclo[3.2.1]octan-3-yl]pyridazin-3-yl]phenol (691 mg, 1.38 mmol, 1 equiv.) in methanol (15 mL) and dichloromethane (5 mL), acetic acid (82 mg, 1.38 mmol, 1 equiv.) was added. The solution was stirred at 30°C for 0.5 h. Sodium cyanoborohydride (129 mg, 2.06 mmol, 1.5 equiv) was then added to the solution and stirred for 7 h at 30° C. The solvent was removed in vacuo and the crude product was purified by preparative HPLC (Column: Phenomenex Synergi Max-RP 250*50 mm*10 um; Mobile phase: [Water (10 mM NH4HCO3)-ACN]; B%: 45ACN%-75ACN%, 28 min, 78% min). (2S,4R)-1-[(2R)-2-[3-[2-[4-[2-[[4-[3-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl]-2-pyridyl]oxy]ethyl]piperazin-1-yl]ethoxy]isoxazol-5-yl]-3-methyl-butanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (840 mg, 0.8 mmol, 58% yield, 98% purity) was obtained as a white solid.
例示的な化合物59および60の例示的な合成
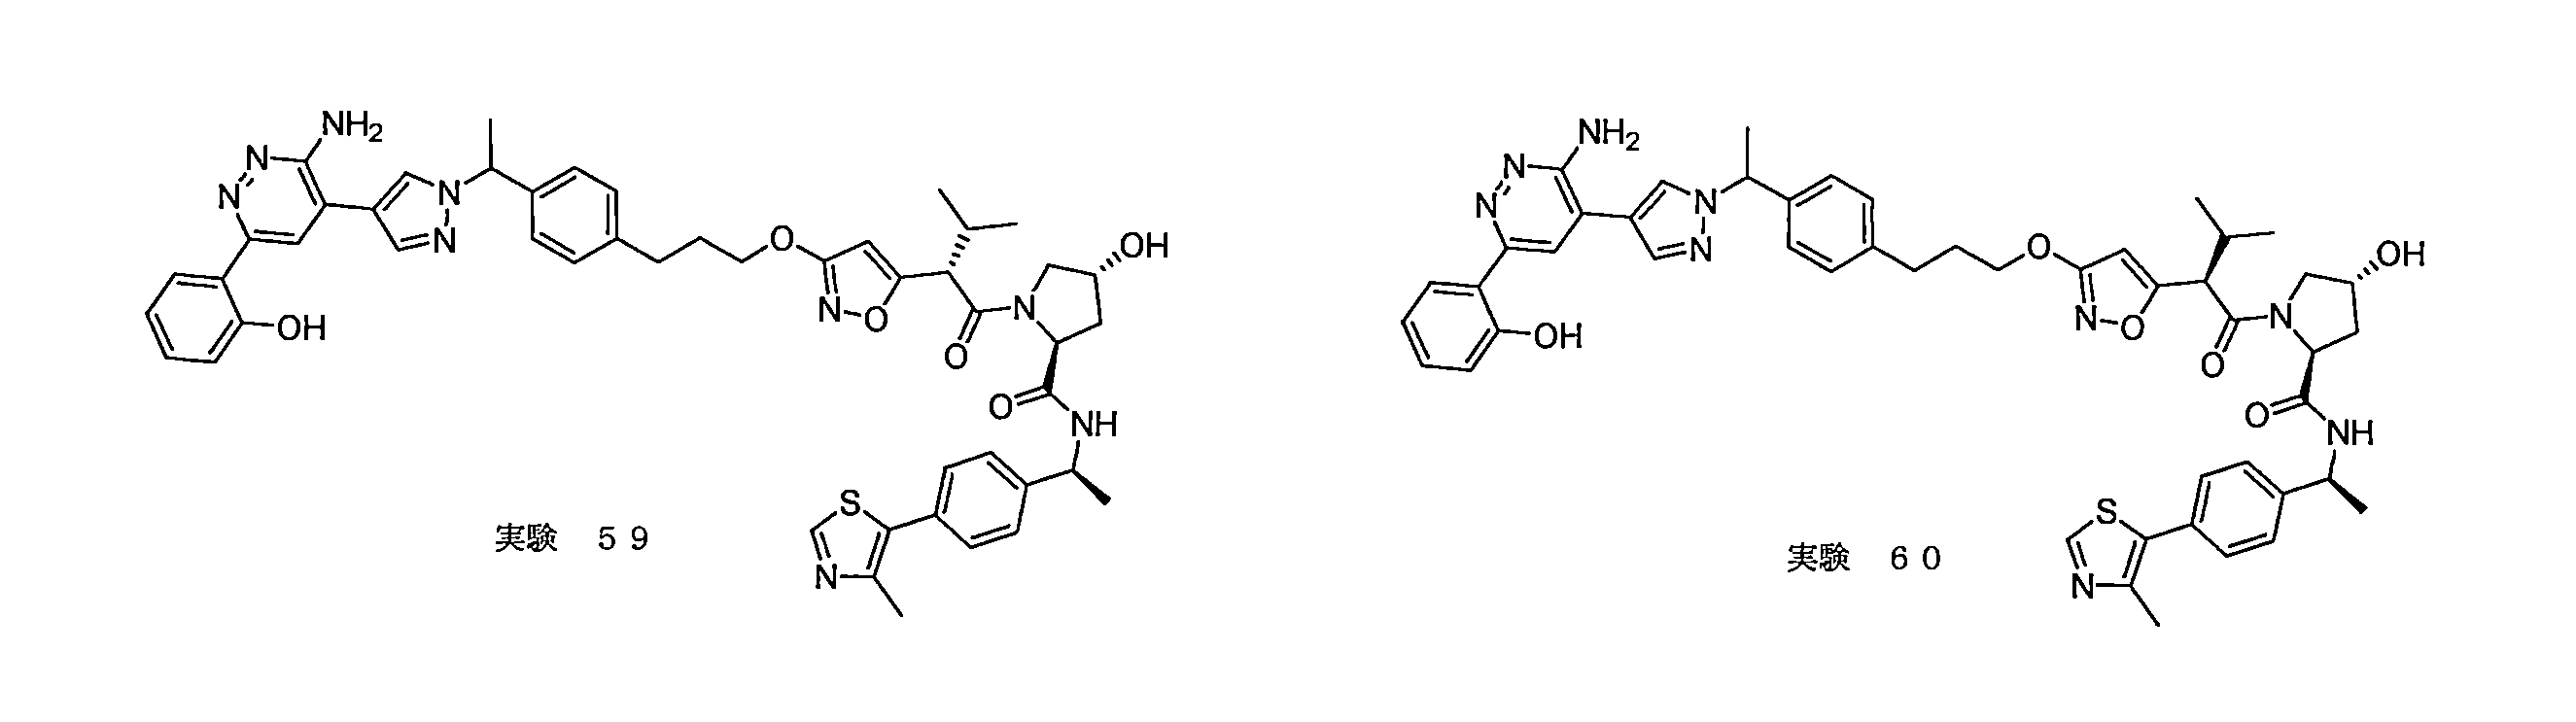
他の実施例について上に記載される手順を使用して、以下のスキームに記載されるように調製した。

例示的な化合物61の例示的な合成

工程1
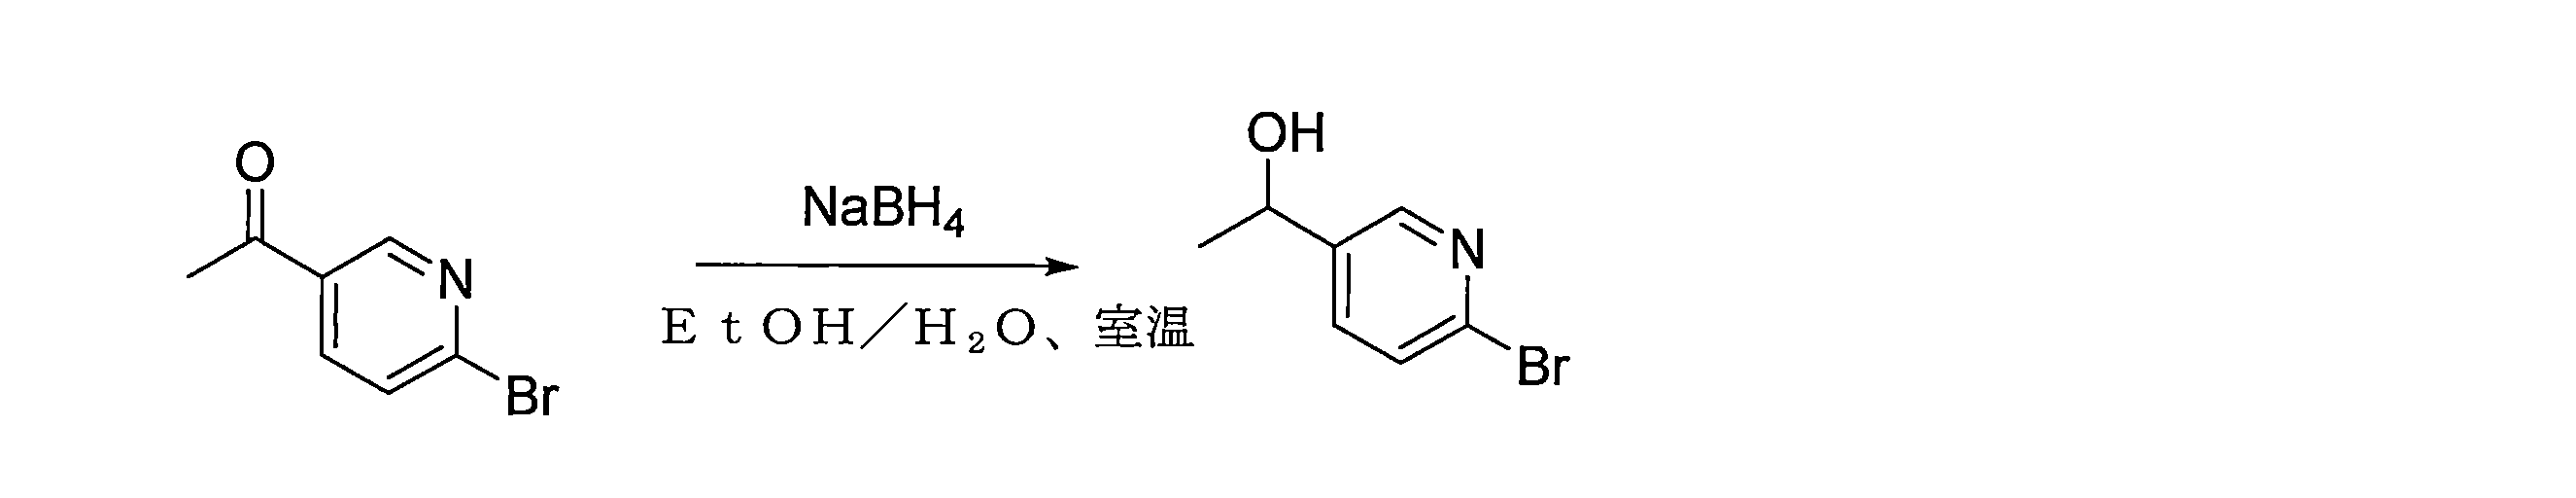
500mLの丸底フラスコに、EtOH(200mL)中の1-(6-ブロモピリジン-3-イル)エタン-1-オン(10g、49.99mmol、1当量)の溶液に、水(80mL)中の水素化ホウ素ナトリウム(4g、105.73mmol、2.11当量)を室温で添加した。得られた混合物を、室温で30分間撹拌した。混合物を、真空下で濃縮した。残渣を、シリカゲルカラムクロマトグラフィーによって精製し、酢酸エチル/石油エーテル(1:1)で溶出して、1-(6-ブロモピリジン-3-イル)エタン-1-オール(9.2g、91%)を黄色の油として得た。 In a 500 mL round bottom flask, to a solution of 1-(6-bromopyridin-3-yl)ethan-1-one (10 g, 49.99 mmol, 1 equiv.) in EtOH (200 mL) was added sodium borohydride (4 g, 105.73 mmol, 2.11 equiv.) in water (80 mL) at room temperature. The resulting mixture was stirred at room temperature for 30 minutes. The mixture was concentrated under vacuum. The residue was purified by silica gel column chromatography and eluted with ethyl acetate/petroleum ether (1:1) to give 1-(6-bromopyridin-3-yl)ethan-1-ol (9.2 g, 91%) as a yellow oil.
工程2

100mLの丸底フラスコに、ジクロロメタン(30mL)中の1-(6-ブロモピリジン-3-イル)エタン-1-オール(2g、9.90mmol、1当量)およびトリエチルアミン(3g、0.03mmol)の溶液に、メタンスルホニルクロリド(1.3g、11.35mmol、1.15当量)を0℃で添加した。得られた混合物を、室温で16時間撹拌した。得られた混合物を、ジクロロメタン(30mL×3)で抽出した。合わせた有機層をブライン(20mL×1)で洗浄し、無水硫酸ナトリウム上で乾燥させた。濾過した後、濾液を減圧下で濃縮した。これにより、1-(6-ブロモピリジン-3-イル)エチルメタンスルホネート(1.6g、23%)を黄色の固体として得た。 In a 100 mL round bottom flask, methanesulfonyl chloride (1.3 g, 11.35 mmol, 1.15 equiv.) was added to a solution of 1-(6-bromopyridin-3-yl)ethan-1-ol (2 g, 9.90 mmol, 1 equiv.) and triethylamine (3 g, 0.03 mmol) in dichloromethane (30 mL) at 0 °C. The resulting mixture was stirred at room temperature for 16 h. The resulting mixture was extracted with dichloromethane (30 mL x 3). The combined organic layers were washed with brine (20 mL x 1) and dried over anhydrous sodium sulfate. After filtration, the filtrate was concentrated under reduced pressure. This gave 1-(6-bromopyridin-3-yl)ethyl methanesulfonate (1.6 g, 23%) as a yellow solid.
工程3

25mLの密封チューブに、1-(6-ブロモピリジン-3-イル)エチルメタンスルホネート(1.4g、1.2当量)、6-[2-(メトキシメトキシ)フェニル]-4-(1H-ピラゾール-4-イル)ピリダジン-3-アミン(350mg、1当量)、および炭酸カリウム(490mg、3.0当量)を、N,N-ジメチルホルムアミド(15mL)に室温で添加した。得られた混合物を、窒素雰囲気下で、60℃で2時間撹拌した。水層をジクロロメタン(50mL×3)で抽出した。残渣を、ブライン(20mL×1)で洗浄し、無水硫酸ナトリウムで乾燥させた。濾過した後、濾液を減圧下で濃縮した。残渣を、シリカゲルカラムクロマトグラフィーによって精製し、ジクロロメタン/メチルアルコール(10:1)で溶出して、4-[1-[1-(6-ブロモピリジン-3-イル)エチル]-1H-ピラゾール-4-イル]-6-[2-(メトキシメトキシ)フェニル]ピリダジン-3-アミン(90mg、16%)を黄色の固体として得た。 In a 25 mL sealed tube, 1-(6-bromopyridin-3-yl)ethyl methanesulfonate (1.4 g, 1.2 eq.), 6-[2-(methoxymethoxy)phenyl]-4-(1H-pyrazol-4-yl)pyridazin-3-amine (350 mg, 1 eq.), and potassium carbonate (490 mg, 3.0 eq.) were added to N,N-dimethylformamide (15 mL) at room temperature. The resulting mixture was stirred at 60° C. for 2 hours under nitrogen atmosphere. The aqueous layer was extracted with dichloromethane (50 mL×3). The residue was washed with brine (20 mL×1) and dried over anhydrous sodium sulfate. After filtration, the filtrate was concentrated under reduced pressure. The residue was purified by silica gel column chromatography and eluted with dichloromethane/methyl alcohol (10:1) to obtain 4-[1-[1-(6-bromopyridin-3-yl)ethyl]-1H-pyrazol-4-yl]-6-[2-(methoxymethoxy)phenyl]pyridazin-3-amine (90 mg, 16%) as a yellow solid.
上記の他の実施例について記載される手順を使用して、以下のスキームに従って、4-[1-[1-(6-ブロモピリジン-3-イル)エチル]-1H-ピラゾール-4-イル]-6-[2-(メトキシメトキシ)フェニル]ピリダジン-3-アミンを最終化合物に変換した。

例示的な化合物62の例示的な合成
工程1

ジメチルホルムアミド(3mL)中の6-[2-(メトキシメトキシ)フェニル]-4-(1H-ピラゾール-4-イル)ピリダジン-3-アミン(300mg、1.01mmol、1当量)、1-ブロモ-2-(2-ブロモエトキシ)エタン(702mg、3.03mmol、3当量)の混合物に、炭酸カリウム(418mg、3.03mmol、3当量)を添加した。次いで、反応混合物を70℃で2時間撹拌した。水(10mL)を添加し、反応混合物を酢酸エチル(20mL×3)で抽出した。合わせた有機層をブライン(15mL×2)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、減圧下で濃縮して、残渣を得た。分取TLC(ジクロロメタン:メタノール=10:1)によって残渣を精製した。4-[1-[2-(2-ブロモエトキシ)エチル]ピラゾール-4-イル]-6-[2-(メトキシメトキシ)フェニル]ピリダジン-3-アミン(100mg、0.22mmol、収率22%)を黄色の油として得た。 To a mixture of 6-[2-(methoxymethoxy)phenyl]-4-(1H-pyrazol-4-yl)pyridazin-3-amine (300 mg, 1.01 mmol, 1 eq.), 1-bromo-2-(2-bromoethoxy)ethane (702 mg, 3.03 mmol, 3 eq.) in dimethylformamide (3 mL), potassium carbonate (418 mg, 3.03 mmol, 3 eq.) was added. The reaction mixture was then stirred at 70°C for 2 h. Water (10 mL) was added and the reaction mixture was extracted with ethyl acetate (20 mL x 3). The combined organic layers were washed with brine (15 mL x 2), dried over anhydrous sodium sulfate, filtered, and concentrated under reduced pressure to give a residue. The residue was purified by preparative TLC (dichloromethane:methanol = 10:1). 4-[1-[2-(2-bromoethoxy)ethyl]pyrazol-4-yl]-6-[2-(methoxymethoxy)phenyl]pyridazin-3-amine (100 mg, 0.22 mmol, 22% yield) was obtained as a yellow oil.
工程2

アセトニトリル(2mL)中の4-[1-[2-(2-ブロモエトキシ)エチル]ピラゾール-4-イル]-6-[2-(メトキシメトキシ)フェニル]ピリダジン-3-アミン(100mg、0.22mmol、1当量)およびtert-ブチルピペラジン-1-カルボキシレート(83mg、0.44mmol、2当量)の溶液に、N,N-ジイソプロピルエチルアミン(86.49mg、0.66mmol、3当量)を添加した。次いで、反応混合物を100℃で12時間撹拌した。水(5mL)を添加し、反応混合物を酢酸エチル(15mL×3)で抽出した。合わせた有機層をブライン(10mL×3)で洗浄し、無水硫酸ナトリウム上で乾燥させ、濾過し、減圧下で濃縮して残渣を得た。分取TLC(ジクロロメタン:メタノール=10:1)によって残渣を精製した。化合物tert-ブチル4-[2-[2-[4-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]ピラゾール-1-イル]エトキシ]エチル]ピペラジン-1-カルボキシレート(93mg、0.15mmol、収率71%)を黄色の固体として得た。 To a solution of 4-[1-[2-(2-bromoethoxy)ethyl]pyrazol-4-yl]-6-[2-(methoxymethoxy)phenyl]pyridazin-3-amine (100 mg, 0.22 mmol, 1 eq.) and tert-butyl piperazine-1-carboxylate (83 mg, 0.44 mmol, 2 eq.) in acetonitrile (2 mL), N,N-diisopropylethylamine (86.49 mg, 0.66 mmol, 3 eq.) was added. The reaction mixture was then stirred at 100°C for 12 h. Water (5 mL) was added and the reaction mixture was extracted with ethyl acetate (15 mL x 3). The combined organic layers were washed with brine (10 mL x 3), dried over anhydrous sodium sulfate, filtered, and concentrated under reduced pressure to give a residue. The residue was purified by preparative TLC (dichloromethane:methanol = 10:1). The compound tert-butyl 4-[2-[2-[4-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]pyrazol-1-yl]ethoxy]ethyl]piperazine-1-carboxylate (93 mg, 0.15 mmol, 71% yield) was obtained as a yellow solid.
上記の他の実施例について記載される手順を使用して、以下のスキームに従って、tert-ブチル4-[2-[2-[4-[3-アミノ-6-[2-(メトキメトキシ)フェニル]ピリダジン-4-イル]ピラゾール-1-イル]エトキシ]エチル]ピペラジン-1-カルボキシレートを最終化合物に変換した。
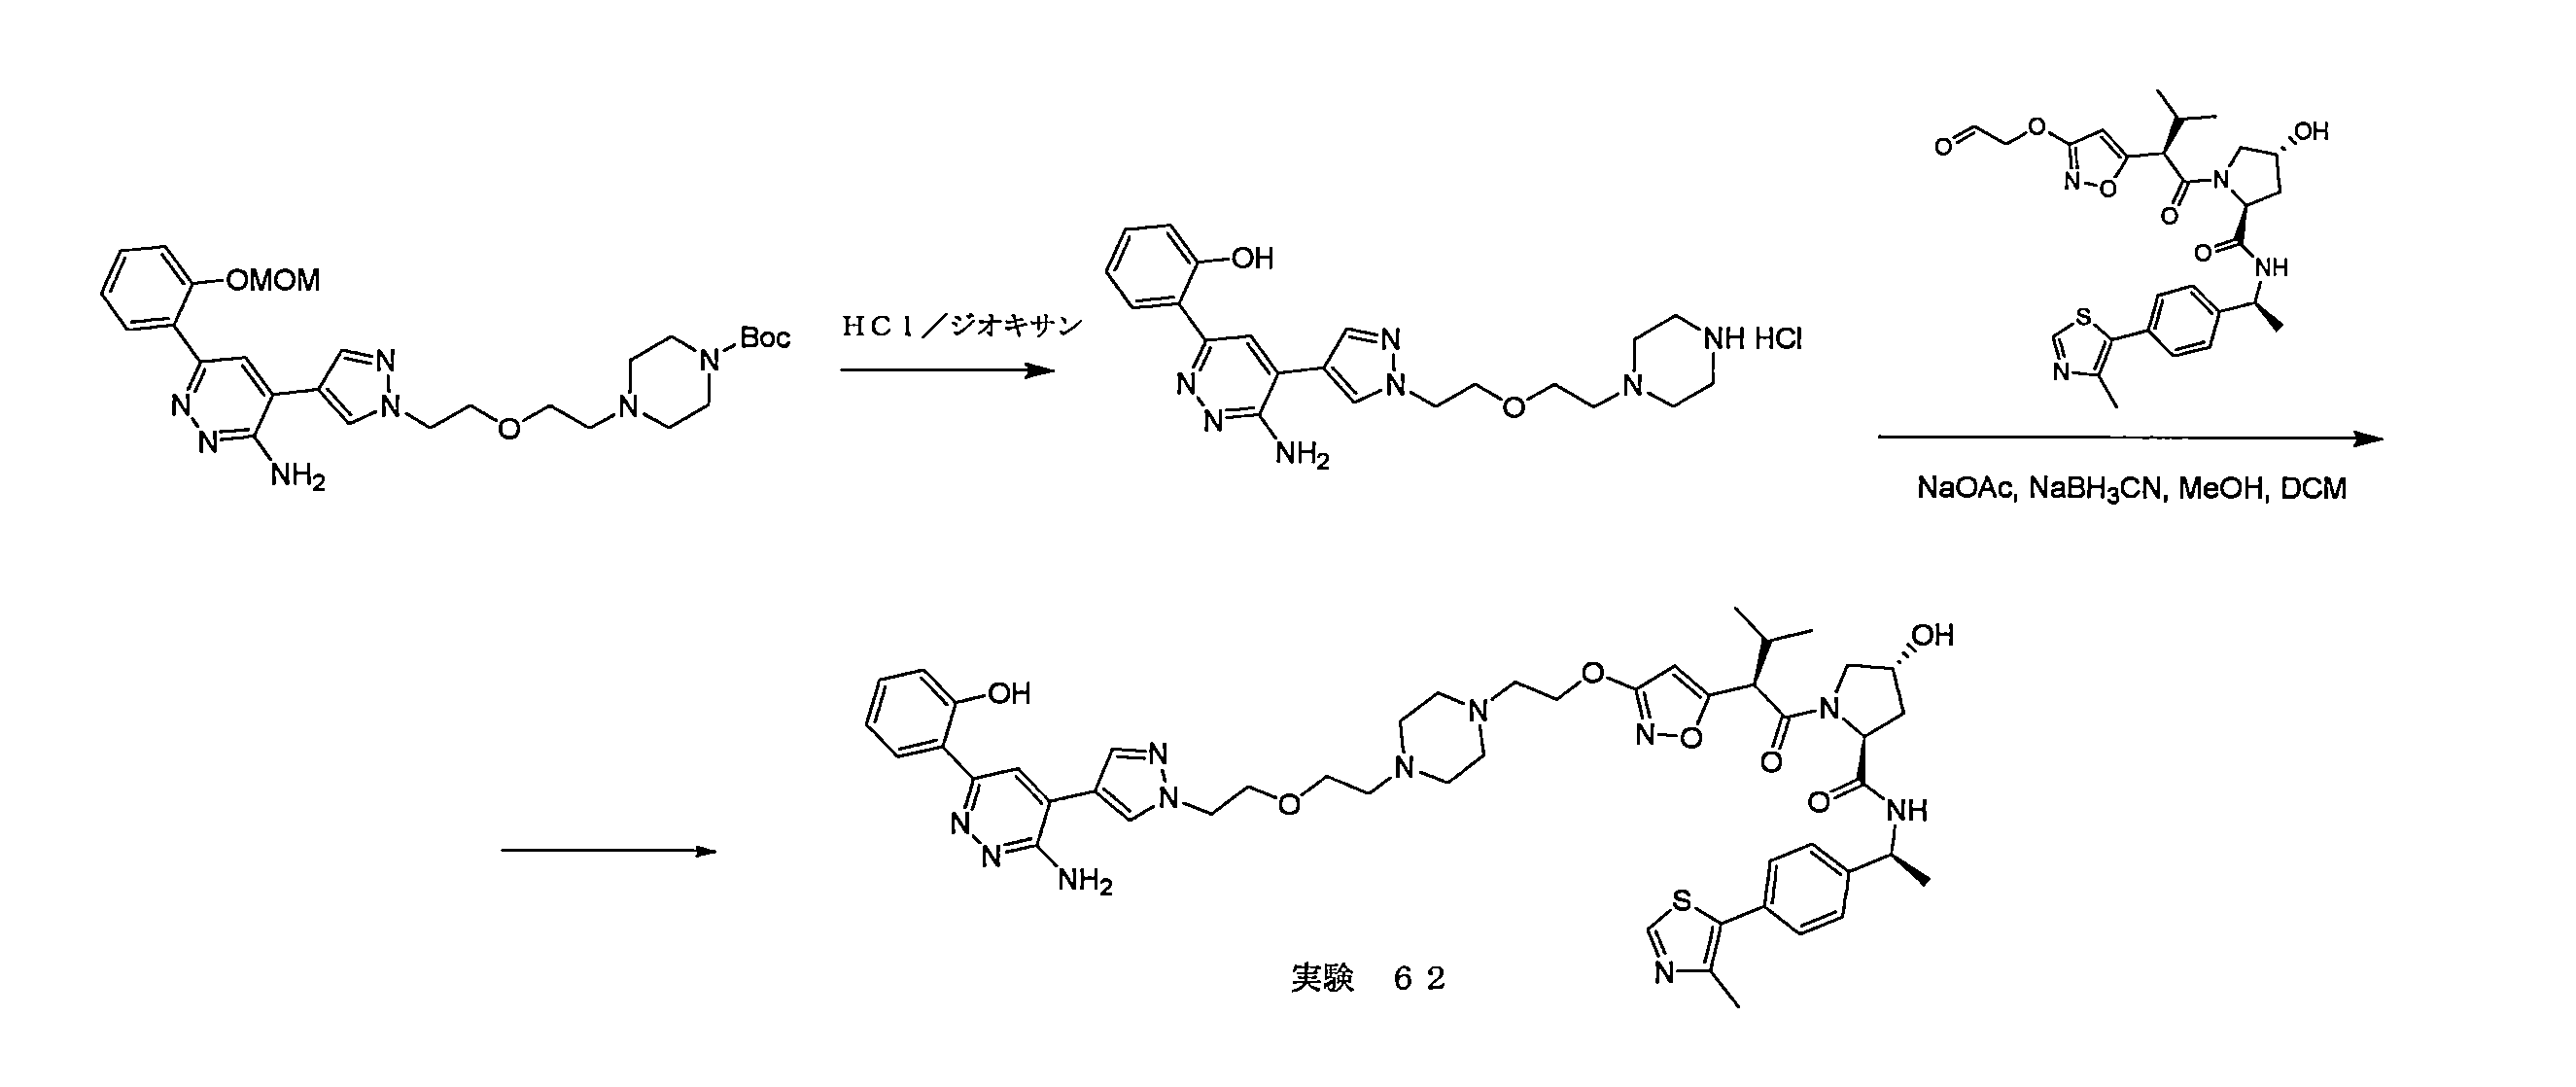
例示的な化合物165および124を、類似の手順を使用して調製した。 Exemplary compounds 165 and 124 were prepared using similar procedures.
例示的な化合物64の例示的な合成
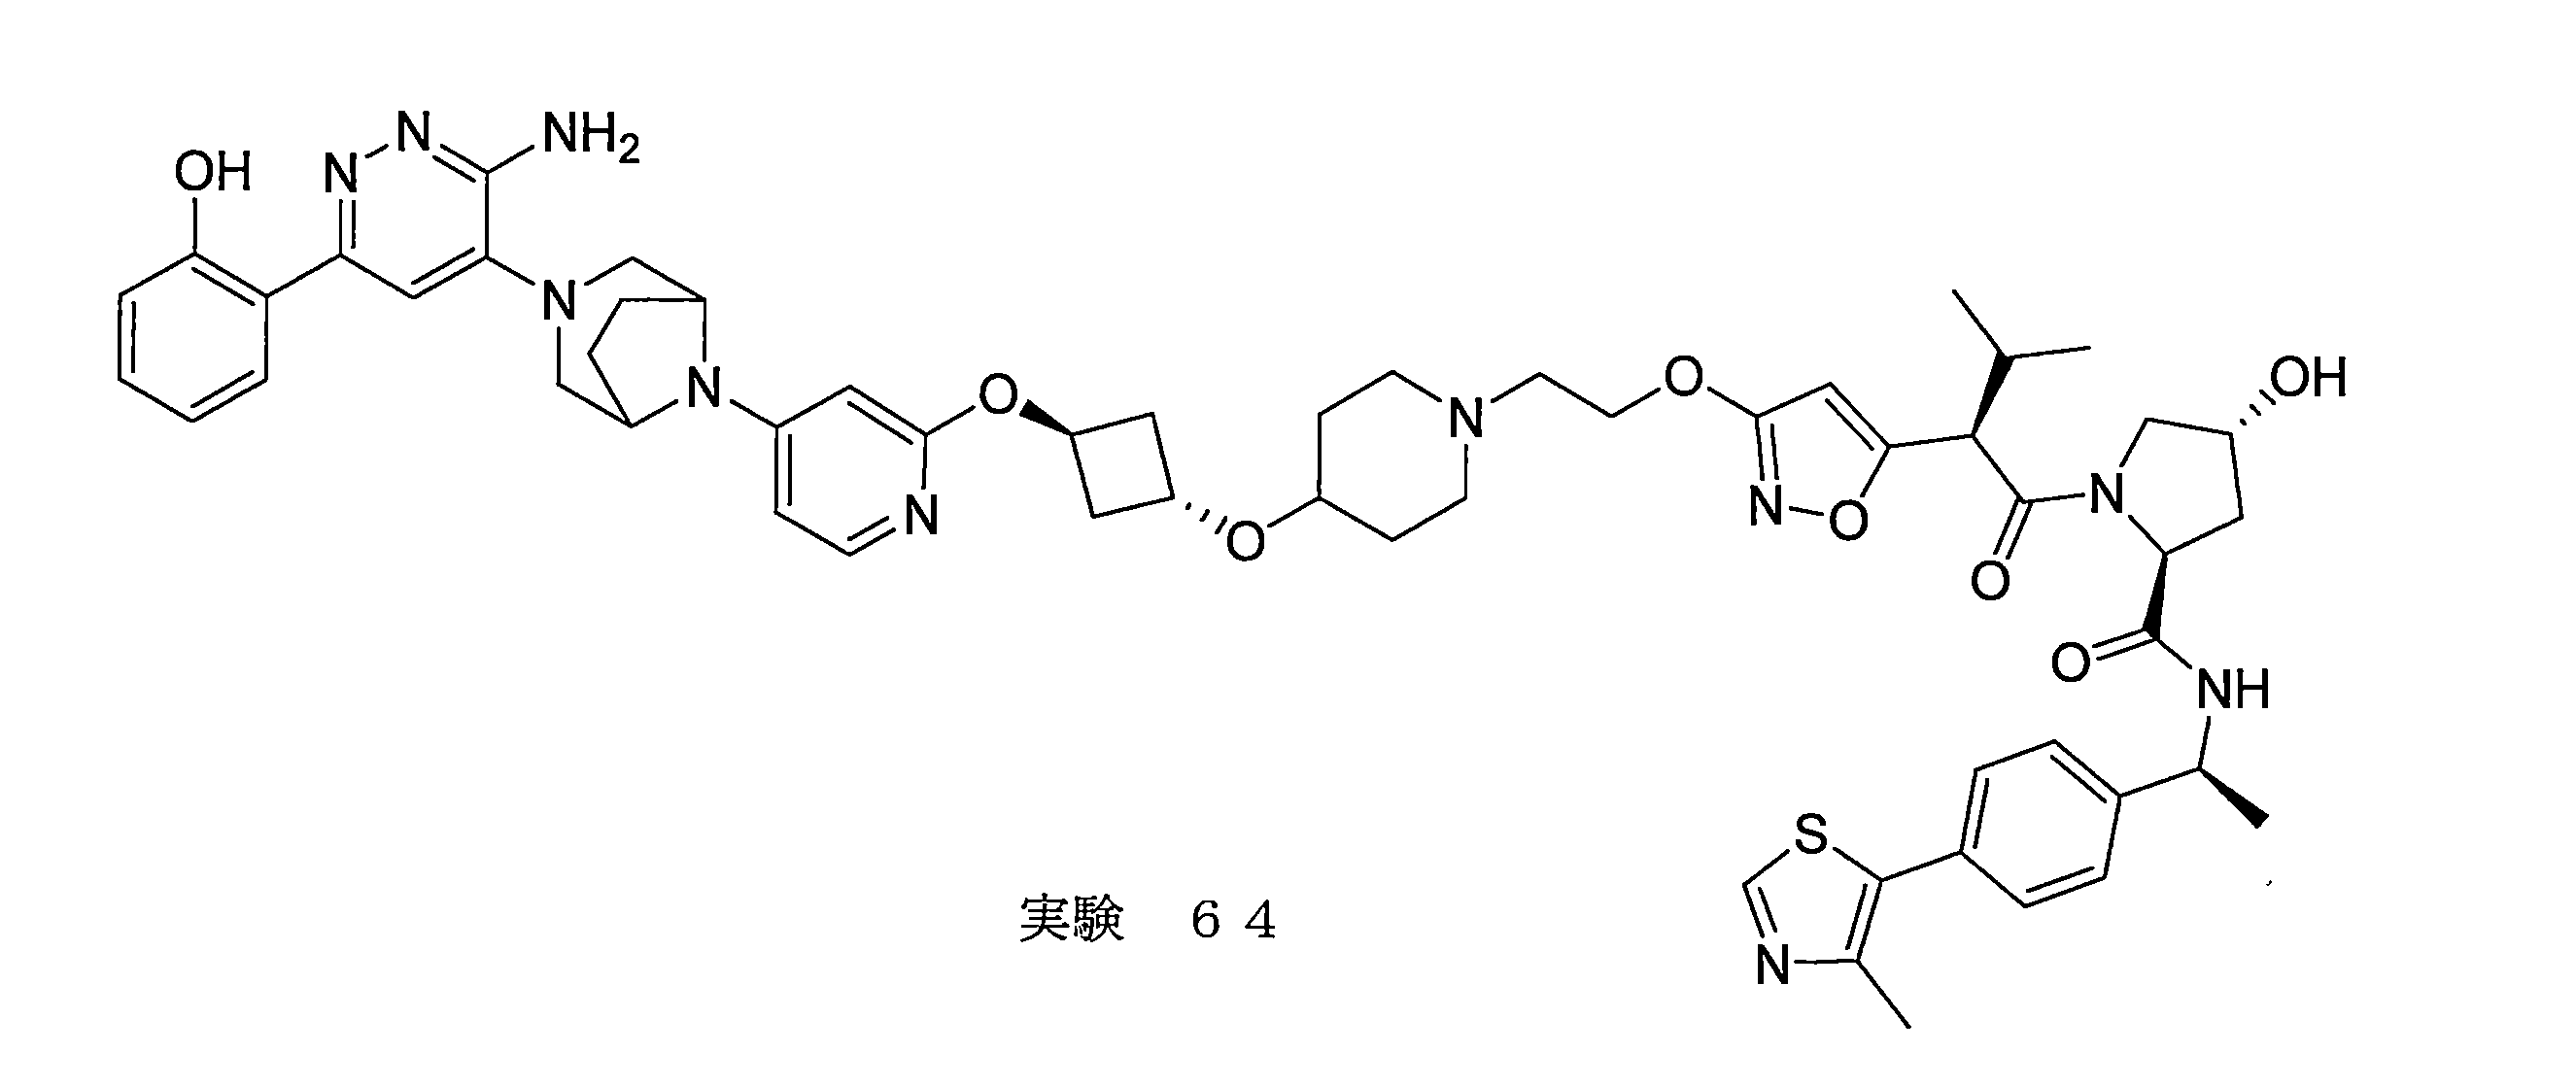
工程1

テトラヒドロフラン(200mL)中のピリジン-4-オール(3.20g、33.66mmol、1.5当量)および3-ベンジルオキシシクロブタノール(4g、22.44mmol、1当量)の溶液に、トリフェニルホスフィン(7.06g、26.93mmol、1.2当量)およびアゾジカルボン酸ジイソプロピル(5.45g、26.93mmol、1.2当量)を、窒素下、10℃で一度に添加した。混合物を50℃で12時間撹拌した。反応混合物を減圧下で濃縮して、テトラヒドロフランを除去した。水(50mL)を混合物に注ぎ、1分間撹拌した。水層をジクロロメタン(50mL×3)で抽出した。合わせた有機層をブライン(50mL×2)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、真空下で濃縮させた。残基をシリカゲルカラムクロマトグラフィー(石油エーテル:テトラヒドロフラン=20:1、5:1)によって精製した。残渣を、フラッシュC18カラムクロマトグラフィー[アセトニトリル:水(0.5%水酸化アンモニウム)=5%~50%]によって精製した。化合物4-(3-ベンジルオキシシクロブトキシ)ピリジン(3.2g、12.53mmol、収率55%)を白色の固体として得た。 To a solution of pyridin-4-ol (3.20 g, 33.66 mmol, 1.5 eq.) and 3-benzyloxycyclobutanol (4 g, 22.44 mmol, 1 eq.) in tetrahydrofuran (200 mL), triphenylphosphine (7.06 g, 26.93 mmol, 1.2 eq.) and diisopropyl azodicarboxylate (5.45 g, 26.93 mmol, 1.2 eq.) were added in one portion under nitrogen at 10 °C. The mixture was stirred at 50 °C for 12 h. The reaction mixture was concentrated under reduced pressure to remove tetrahydrofuran. Water (50 mL) was poured into the mixture and stirred for 1 min. The aqueous layer was extracted with dichloromethane (50 mL x 3). The combined organic layers were washed with brine (50 mL x 2), dried over anhydrous sodium sulfate, filtered, and concentrated under vacuum. The residue was purified by silica gel column chromatography (petroleum ether:tetrahydrofuran = 20:1, 5:1). The residue was purified by flash C18 column chromatography [acetonitrile:water (0.5% ammonium hydroxide) = 5% to 50%]. The compound 4-(3-benzyloxycyclobutoxy)pyridine (3.2 g, 12.53 mmol, yield 55%) was obtained as a white solid.
工程2

トルエン(65mL)中の4-(3-ベンジルオキシシクロブトキシ)ピリジン(4.2g、16.45mmol、1当量)の溶液に、臭化ベンジル(2.81g、16.45mmol、1当量)を添加した。混合物を80℃で12時間撹拌した。反応混合物を減圧下で濃縮して、トルエンを除去した。粗生成物を石油エーテル(80mL)で粉砕した。化合物1-ベンジル-4-(3-ベンジルオキシシクロブトキシ)ピリジン-1-臭化イウム(6.5g、15.25mmol、収率92%)を白色の固体として得た。 To a solution of 4-(3-benzyloxycyclobutoxy)pyridine (4.2 g, 16.45 mmol, 1 equiv.) in toluene (65 mL) was added benzyl bromide (2.81 g, 16.45 mmol, 1 equiv.). The mixture was stirred at 80 °C for 12 h. The reaction mixture was concentrated under reduced pressure to remove toluene. The crude product was triturated with petroleum ether (80 mL). The compound 1-benzyl-4-(3-benzyloxycyclobutoxy)pyridine-1-ium bromide (6.5 g, 15.25 mmol, 92% yield) was obtained as a white solid.
工程3
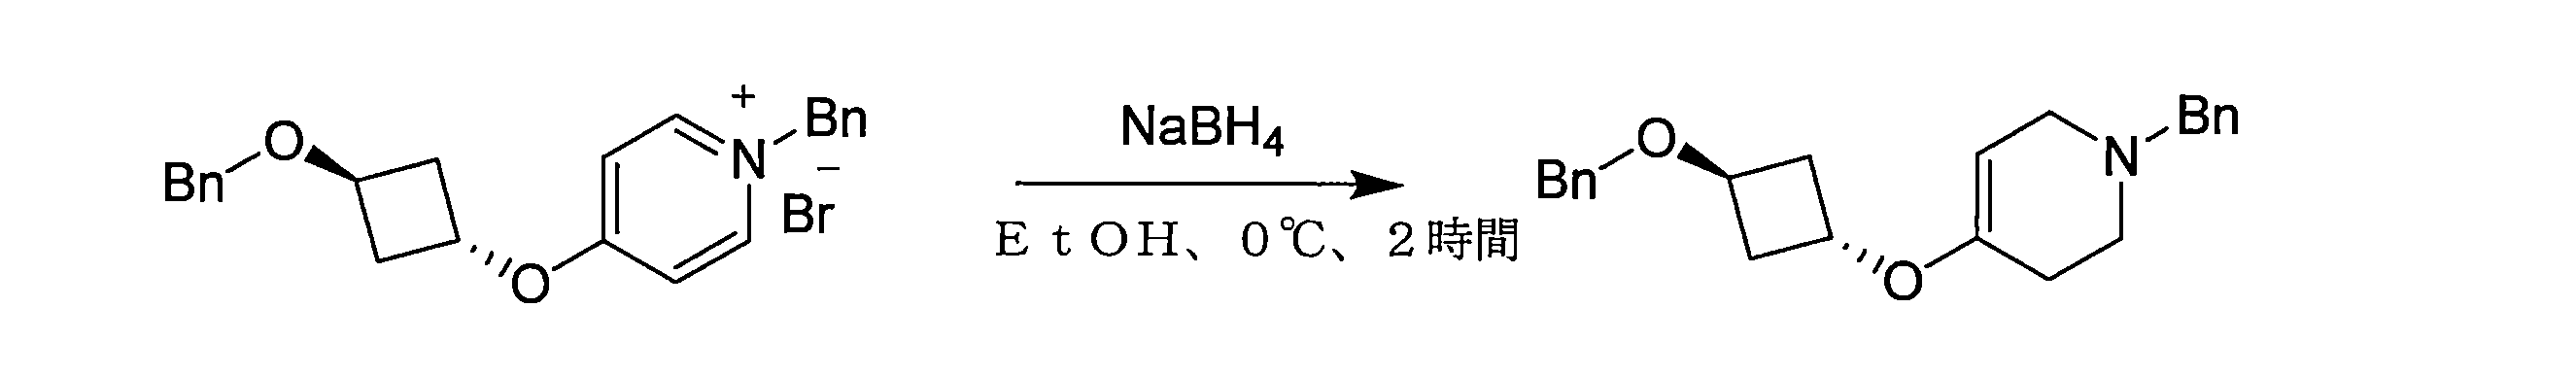
エタノール(120mL)中の1-ベンジル-4-(3-ベンジルオキシシクロブトキシ)ピリジン-1-イウムブロミド(6.5g、15.25mmol、1当量)の溶液に、水素化ホウ素ナトリウム(3.46g、91.47mmol、6当量)を0℃で添加した。混合物を15℃で4時間撹拌した。反応混合物を減圧下で濃縮して、エタノールを除去した。残渣を水(25mL)で希釈し、酢酸エチル(50mL×2)で抽出した。合わせた有機層を飽和ブライン(40mL×3)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、真空中で濃縮した。化合物1-ベンジル-4-(3-ベンジルオキシシクロブトキシ)-3,6-ジヒドロ-2H-ピリジン(4.5g、12.88mmol、収率84%)を無色の油として得た。 To a solution of 1-benzyl-4-(3-benzyloxycyclobutoxy)pyridin-1-ium bromide (6.5 g, 15.25 mmol, 1 equiv.) in ethanol (120 mL) was added sodium borohydride (3.46 g, 91.47 mmol, 6 equiv.) at 0°C. The mixture was stirred at 15°C for 4 h. The reaction mixture was concentrated under reduced pressure to remove ethanol. The residue was diluted with water (25 mL) and extracted with ethyl acetate (50 mL x 2). The combined organic layers were washed with saturated brine (40 mL x 3), dried over anhydrous sodium sulfate, filtered and concentrated in vacuum. Compound 1-benzyl-4-(3-benzyloxycyclobutoxy)-3,6-dihydro-2H-pyridine (4.5 g, 12.88 mmol, 84% yield) was obtained as a colorless oil.
工程4

テトラヒドロフラン(95mL)およびエタノール(70mL)中の1-ベンジル-4-(3-ベンジルオキシシクロブトキシ)-3,6-ジヒドロ-2H-ピリジン(4.5g、12.88mmol、1当量)の溶液に、パラジウム活性炭素触媒(0.5g、純度10%)を窒素雰囲気下で添加した。懸濁液を脱気し、水素で3回パージした。混合物を、水素(50Psi)下で、25℃で24時間撹拌し、次いで、35℃で12時間撹拌した。反応混合物を濾過し、濾液を濃縮した。残渣を、シリカゲルカラムクロマトグラフィー(ジクロロメタン:メタノール:水酸化アンモニウム=20:1:0、10:1:0.1)によって精製した。化合物3-[(1-ベンジル-4-ピペリジル)オキシ]シクロブタノール(2.8g、10.71mmol、収率83%)を、無色の油として得た。 To a solution of 1-benzyl-4-(3-benzyloxycyclobutoxy)-3,6-dihydro-2H-pyridine (4.5 g, 12.88 mmol, 1 equiv.) in tetrahydrofuran (95 mL) and ethanol (70 mL), palladium-activated carbon catalyst (0.5 g, 10% purity) was added under nitrogen atmosphere. The suspension was degassed and purged with hydrogen three times. The mixture was stirred under hydrogen (50 Psi) at 25°C for 24 h and then at 35°C for 12 h. The reaction mixture was filtered and the filtrate was concentrated. The residue was purified by silica gel column chromatography (dichloromethane:methanol:ammonium hydroxide=20:1:0, 10:1:0.1). Compound 3-[(1-benzyl-4-piperidyl)oxy]cyclobutanol (2.8 g, 10.71 mmol, 83% yield) was obtained as a colorless oil.
工程5
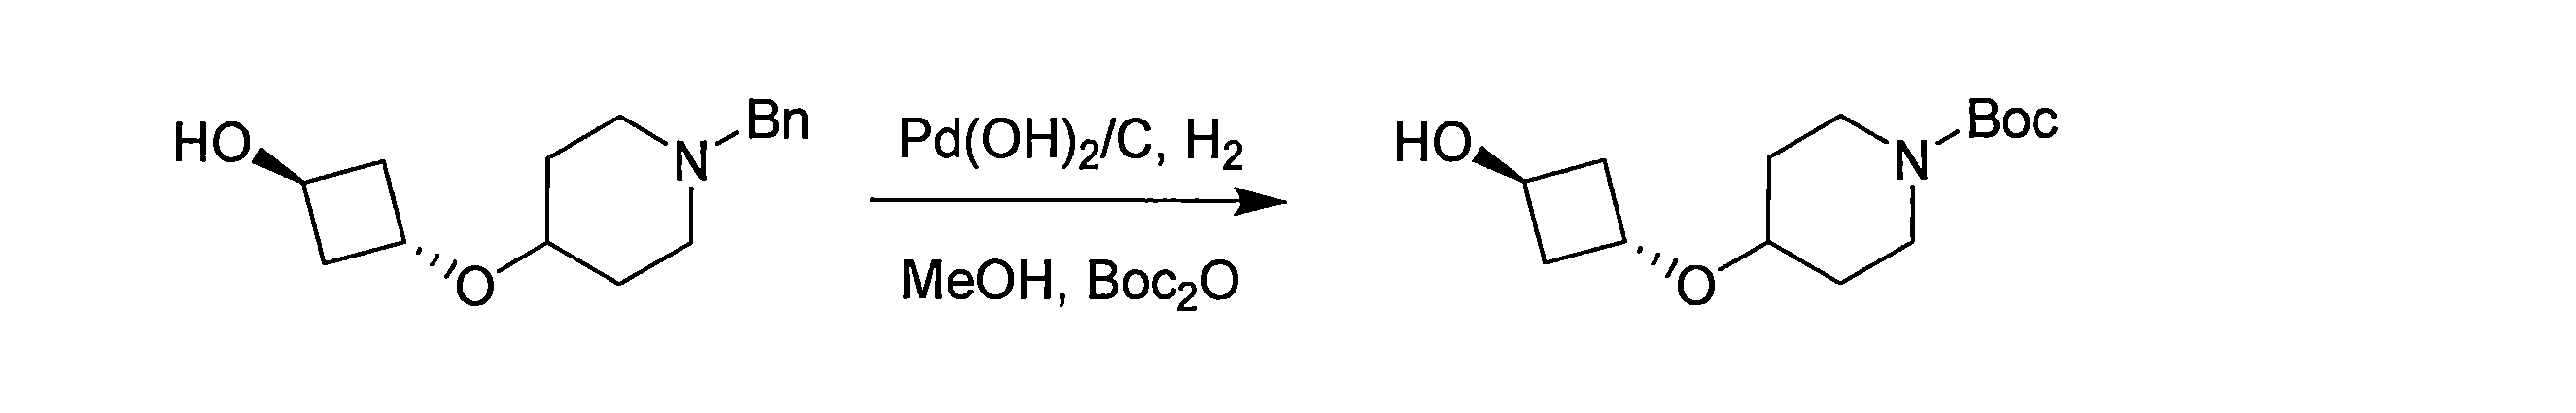
メタノール(10mL)中の3-[(1-ベンジル-4-ピペリジル)オキシ]シクロブタノール(1.1g、4.21mmol、1当量)の溶液に、水酸化パラジウム(591mg)およびジ-tert-ブチルジカーボネートエステル(1.84g、8.42mmol、2当量)を窒素雰囲気下で添加した。懸濁液を脱気し、水素で3回パージした。混合物を、水素(50psi)下で、25℃で12時間撹拌した。反応混合物を濾過し、フィルターを濃縮した。残渣を、シリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=20:1~2:1)によって精製した。化合物tert-ブチル4-(3-ヒドロキシシクロブトキシ)ピペリジン-1-カルボキシレート(820mg、3.02mmol、収率71%)を無色の油として得た。 To a solution of 3-[(1-benzyl-4-piperidyl)oxy]cyclobutanol (1.1 g, 4.21 mmol, 1 equiv.) in methanol (10 mL), palladium hydroxide (591 mg) and di-tert-butyl dicarbonate ester (1.84 g, 8.42 mmol, 2 equiv.) were added under nitrogen atmosphere. The suspension was degassed and purged with hydrogen three times. The mixture was stirred under hydrogen (50 psi) at 25°C for 12 h. The reaction mixture was filtered and the filter was concentrated. The residue was purified by silica gel chromatography (petroleum ether:ethyl acetate=20:1-2:1). The compound tert-butyl 4-(3-hydroxycyclobutoxy)piperidine-1-carboxylate (820 mg, 3.02 mmol, 71% yield) was obtained as a colorless oil.
工程6

アセトニトリル(10mL)中tert-ブチル4-(3-ヒドロキシシクロブトキシ)ピペリジン-1-カルボキシレート(1g、3.69mmol、1当量)、4-ブロモ-2-フルオロ-ピリジン(778mg、4.42mmol、1.2当量)、炭酸セシウム(2.40g、7.37mmol、2当量)の混合物を脱気し、窒素で3回パージし、次いで、混合物を窒素雰囲気下、90℃で12時間撹拌した。反応混合物を濾過し、濃縮して、粗生成物を得た。粗生成物をシリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=1:0~10:1)によって精製した。化合物tert-ブチル4-[3-[(4-ブロモ-2-ピリジル)オキシ]シクロブトキシ]ピペリジン-1-カルボキシレート(1.36g、3.18mmol、収率86%)を無色の油として得た。 A mixture of tert-butyl 4-(3-hydroxycyclobutoxy)piperidine-1-carboxylate (1 g, 3.69 mmol, 1 eq.), 4-bromo-2-fluoro-pyridine (778 mg, 4.42 mmol, 1.2 eq.), and cesium carbonate (2.40 g, 7.37 mmol, 2 eq.) in acetonitrile (10 mL) was degassed and purged with nitrogen three times, and then the mixture was stirred at 90° C. under nitrogen atmosphere for 12 h. The reaction mixture was filtered and concentrated to give the crude product. The crude product was purified by silica gel chromatography (petroleum ether:ethyl acetate=1:0-10:1). The compound tert-butyl 4-[3-[(4-bromo-2-pyridyl)oxy]cyclobutoxy]piperidine-1-carboxylate (1.36 g, 3.18 mmol, 86% yield) was obtained as a colorless oil.
工程7

ジクロロメタン(140mL)中のtert-ブチル3,8-ジアザビシクロ[3.2.1]オクタン-8-カルボキシレート(10g、47.11mmol、1当量)の溶液に、CbzCl(9.64g、56.53mmol、8.04mL、1.2当量)およびトリエチルアミン(8.58g、84.79mmol、11.80mL、1.8当量)を添加した。混合物を20℃で20時間撹拌した。TLC(ジクロロメタン:メタノール=10:1)は、反応物1が完全に消費されていないことを示したため、CbzCl(1.61g、9.42mmol、1.34mL、0.2当量)を再び添加した。反応混合物を20℃で2時間撹拌した。反応混合物を水(80mL)で2回洗浄し、有機層を無水硫酸ナトリウムで乾燥させ、濃縮して、粗生成物を得た。粗生成物をシリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=1:0~4:1)によって精製した。化合物8-ベンジル3-(tert-ブチル)3,8-ジアザビシクロ[3.2.1]オクタン-3,8-ジカルボキシレート(11.48g、33.14mmol、収率70%)を無色の油として得た。 To a solution of tert-butyl 3,8-diazabicyclo[3.2.1]octane-8-carboxylate (10 g, 47.11 mmol, 1 eq.) in dichloromethane (140 mL), CbzCl (9.64 g, 56.53 mmol, 8.04 mL, 1.2 eq.) and triethylamine (8.58 g, 84.79 mmol, 11.80 mL, 1.8 eq.) were added. The mixture was stirred at 20 °C for 20 h. TLC (dichloromethane:methanol = 10:1) showed that reactant 1 was not completely consumed, so CbzCl (1.61 g, 9.42 mmol, 1.34 mL, 0.2 eq.) was added again. The reaction mixture was stirred at 20 °C for 2 h. The reaction mixture was washed twice with water (80 mL), and the organic layer was dried over anhydrous sodium sulfate and concentrated to give the crude product. The crude product was purified by silica gel chromatography (petroleum ether:ethyl acetate=1:0 to 4:1). Compound 8-benzyl 3-(tert-butyl) 3,8-diazabicyclo[3.2.1]octane-3,8-dicarboxylate (11.48 g, 33.14 mmol, 70% yield) was obtained as a colorless oil.
工程8
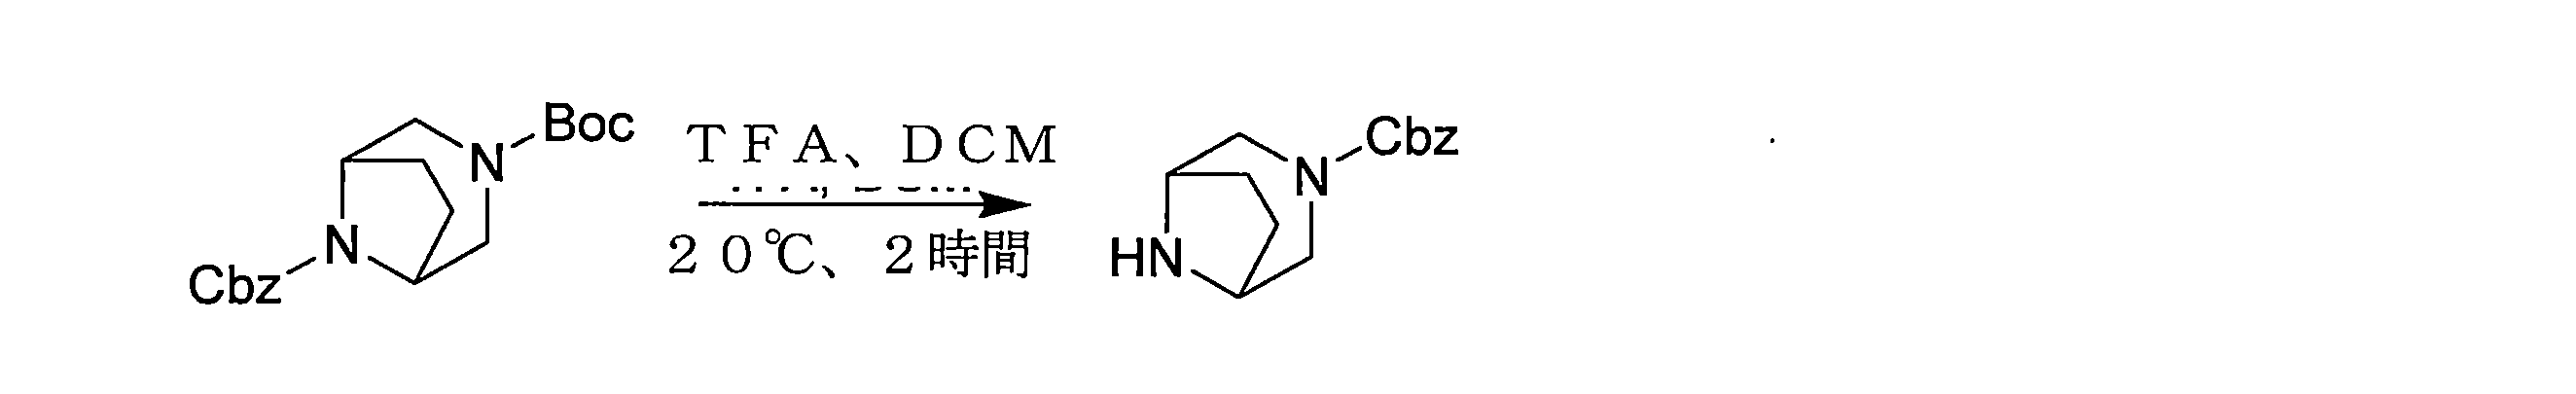
ジクロロメタン(80mL)中の8-ベンジル3-(tert-ブチル)3,8-ジアザビシクロ[3.2.1]オクタン-3,8-ジカルボキシレート(6.20g、17.90mmol、1当量)の溶液に、トリフルオロ酢酸(12.32g、108.05mmol、8.00mL、6.04当量)を添加した。混合物を20℃で2時間撹拌した。反応混合物を飽和重炭酸ナトリウム水溶液でpH=8まで調整し、クロロホルムおよびイソプロパノール(80mL×3、5:1)で抽出した。合わせた層を無水硫酸ナトリウムで乾燥させ、濃縮して、ベンジル3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレート(4.7g、17.68mmol、収率98%、純度92%)を黄色の油として得た。 To a solution of 8-benzyl 3-(tert-butyl) 3,8-diazabicyclo[3.2.1]octane-3,8-dicarboxylate (6.20 g, 17.90 mmol, 1 equiv.) in dichloromethane (80 mL), trifluoroacetic acid (12.32 g, 108.05 mmol, 8.00 mL, 6.04 equiv.) was added. The mixture was stirred at 20 °C for 2 h. The reaction mixture was adjusted to pH = 8 with saturated aqueous sodium bicarbonate and extracted with chloroform and isopropanol (80 mL x 3, 5:1). The combined layers were dried over anhydrous sodium sulfate and concentrated to give benzyl 3,8-diazabicyclo[3.2.1]octane-3-carboxylate (4.7 g, 17.68 mmol, 98% yield, 92% purity) as a yellow oil.
工程9
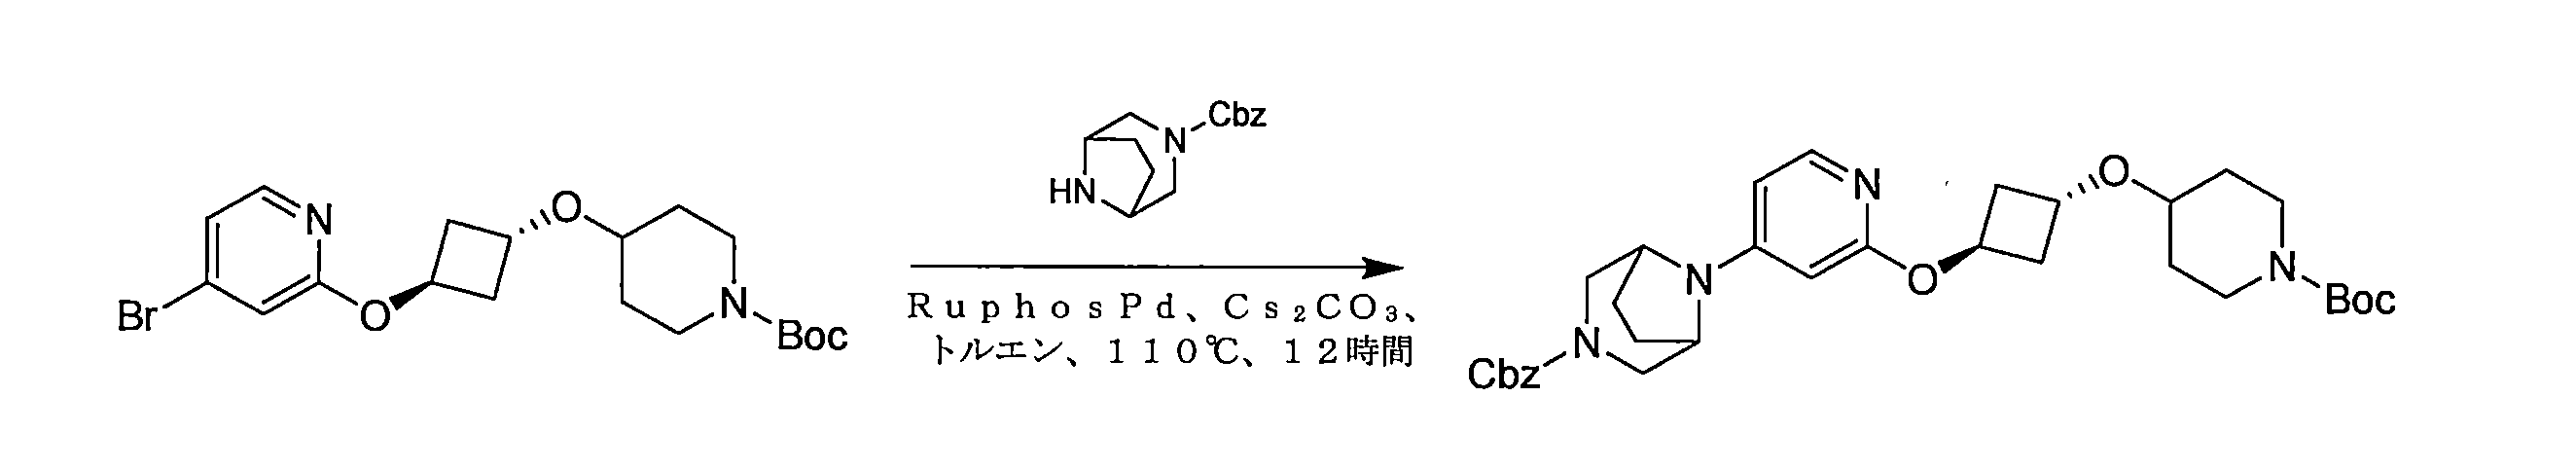
トルエン(27mL)中のベンジル3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレート(784mg、3.18mmol、1当量)、tert-ブチル4-[3-[(4-ブロモ-2-ピリジル)オキシ]シクロブトキシ]ピペリジン-1-カルボキシレート(1.36g、3.18mmol、1当量)、[2-(2-アミノフェニル)フェニル]-クロロパラジウム;ジシクロヘキシル-[2-(2,6-ジイソプロポキシフェニル)フェニル]ホスファン(148mg、0.19mmol、0.06当量)、および炭酸セシウム(2.07g、6.37mmol、2当量)の混合物を脱気し、窒素で3回パージし、次いで、混合物を窒素雰囲気下で、110℃で12時間撹拌した。反応混合物を水(30mL)で希釈し、酢酸エチル(40mL)で2回抽出した。合わせた有機層を飽和塩化ナトリウム水溶液(30mL)で2回洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、減圧下で濃縮して、粗生成物を得た。粗生成物をシリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=1:0~1:1)によって精製した。化合物8-[2-[3-[(1-tert-ブトキシカルボニル-4-ピペリジル)オキシ]シクロブトキシ]-4-ピリジル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレート(1.35g、2.28mmol、収率71%)を無色の油として得た。 A mixture of benzyl 3,8-diazabicyclo[3.2.1]octane-3-carboxylate (784 mg, 3.18 mmol, 1 equiv.), tert-butyl 4-[3-[(4-bromo-2-pyridyl)oxy]cyclobutoxy]piperidine-1-carboxylate (1.36 g, 3.18 mmol, 1 equiv.), [2-(2-aminophenyl)phenyl]-chloropalladium; dicyclohexyl-[2-(2,6-diisopropoxyphenyl)phenyl]phosphane (148 mg, 0.19 mmol, 0.06 equiv.), and cesium carbonate (2.07 g, 6.37 mmol, 2 equiv.) in toluene (27 mL) was degassed and purged with nitrogen three times, then the mixture was stirred under nitrogen atmosphere at 110 °C for 12 h. The reaction mixture was diluted with water (30 mL) and extracted twice with ethyl acetate (40 mL). The combined organic layers were washed twice with saturated aqueous sodium chloride solution (30 mL), dried over anhydrous sodium sulfate, filtered, and concentrated under reduced pressure to obtain the crude product. The crude product was purified by silica gel chromatography (petroleum ether:ethyl acetate=1:0 to 1:1). Compound 8-[2-[3-[(1-tert-butoxycarbonyl-4-piperidyl)oxy]cyclobutoxy]-4-pyridyl]-3,8-diazabicyclo[3.2.1]octane-3-carboxylate (1.35 g, 2.28 mmol, 71% yield) was obtained as a colorless oil.
工程10

テトラヒドロフラン(27mL)およびエタノール(27mL)の混合物中のベンジル8-[2-[3-[(1-tert-ブトキシカルボニル-4-ピペリジル)オキシ]シクロブトキシ]-4-ピリジル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレート(1.35g、2.28mmol、1当量)の溶液に、水酸化パラジウム活性炭素触媒(320mg、純度10%)を窒素下で添加した。懸濁液を真空下で脱気し、水素で数回パージした。混合物を、水素(50psi)下で、60℃で12時間撹拌した。反応混合物を濾過し、濾液を濃縮した。化合物tert-ブチル4-[3-[[4-(3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)-2-ピリジル]オキシ]シクロブトキシ]ピペリジン-1-カルボキシレート(932mg、2.03mmol、収率89%)を茶色の固体として得た。 To a solution of benzyl 8-[2-[3-[(1-tert-butoxycarbonyl-4-piperidyl)oxy]cyclobutoxy]-4-pyridyl]-3,8-diazabicyclo[3.2.1]octane-3-carboxylate (1.35 g, 2.28 mmol, 1 equiv.) in a mixture of tetrahydrofuran (27 mL) and ethanol (27 mL), palladium hydroxide on activated carbon catalyst (320 mg, 10% purity) was added under nitrogen. The suspension was degassed under vacuum and purged several times with hydrogen. The mixture was stirred under hydrogen (50 psi) at 60 °C for 12 h. The reaction mixture was filtered and the filtrate was concentrated. The compound tert-butyl 4-[3-[[4-(3,8-diazabicyclo[3.2.1]octan-8-yl)-2-pyridyl]oxy]cyclobutoxy]piperidine-1-carboxylate (932 mg, 2.03 mmol, 89% yield) was obtained as a brown solid.
工程11

ジメチルスルホキシド(30mL)中のtert-ブチル4-[3-[[4-(3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)-2-ピリジル]オキシ]シクロブトキシ]ピペリジン-1-カルボキシレート(932mg、2.03mmol、1当量)、4-ブロモ-6-クロロ-ピリダジン-3-アミン(508mg、2.44mmol、1.2当量)、N,N-ジイソプロピルエチルアミン(2.63g、20.32mmol、3.54mL、10当量)の混合物を脱気し、窒素で3回パージし、次いで、混合物を窒素雰囲気下で、130℃で3時間撹拌した。反応混合物を水(30mL)で希釈し、酢酸エチル(30mL)で2回抽出した。合わせた有機層を飽和塩化ナトリウム水溶液(30mL)で2回洗浄し、無水硫酸ナトリウムで乾燥させ、濃縮して、粗生成物を得た。残渣を半分取逆相HPLC(カラム:[Phenomenex Gemini C18 250*50 10um;移動相:[水(0.05%水酸化アンモニアv/v)-ACN];B%:45%~70%、24分;53%分)によって精製した。化合物tert-ブチル4-[3-[[4-[3-(3-アミノ-6-クロロ-ピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]-2-ピリジル]オキシ]シクロブトキシ]ピペリジン-1-カルボキシレート(772mg、1.32mmol、収率64%)を黄色の油として得た。 A mixture of tert-butyl 4-[3-[[4-(3,8-diazabicyclo[3.2.1]octan-8-yl)-2-pyridyl]oxy]cyclobutoxy]piperidine-1-carboxylate (932 mg, 2.03 mmol, 1 eq.), 4-bromo-6-chloro-pyridazin-3-amine (508 mg, 2.44 mmol, 1.2 eq.), N,N-diisopropylethylamine (2.63 g, 20.32 mmol, 3.54 mL, 10 eq.) in dimethylsulfoxide (30 mL) was degassed and purged with nitrogen three times, then the mixture was stirred under nitrogen atmosphere at 130 °C for 3 h. The reaction mixture was diluted with water (30 mL) and extracted twice with ethyl acetate (30 mL). The combined organic layers were washed twice with saturated aqueous sodium chloride solution (30 mL), dried over anhydrous sodium sulfate, and concentrated to give the crude product. The residue was purified by semi-preparative reverse phase HPLC (column: [Phenomenex Gemini C18 250*50 10um; mobile phase: [water (0.05% ammonia hydroxide v/v)-ACN]; B%: 45%-70%, 24 min; 53% min). The compound tert-butyl 4-[3-[[4-[3-(3-amino-6-chloro-pyridazin-4-yl)-3,8-diazabicyclo[3.2.1]octan-8-yl]-2-pyridyl]oxy]cyclobutoxy]piperidine-1-carboxylate (772 mg, 1.32 mmol, 64% yield) was obtained as a yellow oil.
工程12

ジオキサン(12mL)および水(2mL)の混合物中のtert-ブチル4-[3-[[4-[3-(3-アミノ-6-クロロ-ピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]-2-ピリジル]オキシ]シクロブトキシ]ピペリジン-1-カルボキシレート(772mg、1.32mmol、1当量)、(2-ヒドロキシフェニル)ボロン酸(218mg、1.58mmol、1.2当量)、テトラキス[トリフェニルホスフィン]パラジウム(0)(152mg、0.13mmol、0.1当量)、炭酸カリウム(364mg、2.63mmol、2当量)の混合物を脱気し、窒素で3回パージし、次いで、混合物を窒素雰囲気下で、90℃で10時間撹拌した。反応混合物を濃縮して、粗生成物を得た。残渣を半分取逆相HPLC(カラム:[Kromasil 250*50mm*10um;移動相:[水(0.1%トリフルオロ酢酸)-ACN];B%:20ACN%~50ACN%、20分)によって精製した。化合物tert-ブチル-4-[3-[[4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]-2-ピリジル]オキシ]シクロブトキシ]ピペリジン-1-カルボキシレート(500mg、0.71mmol、収率54%、純度91%)を黄色の固体として得た。 A mixture of tert-butyl 4-[3-[[4-[3-(3-amino-6-chloro-pyridazin-4-yl)-3,8-diazabicyclo[3.2.1]octan-8-yl]-2-pyridyl]oxy]cyclobutoxy]piperidine-1-carboxylate (772 mg, 1.32 mmol, 1 eq.), (2-hydroxyphenyl)boronic acid (218 mg, 1.58 mmol, 1.2 eq.), tetrakis[triphenylphosphine]palladium(0) (152 mg, 0.13 mmol, 0.1 eq.), potassium carbonate (364 mg, 2.63 mmol, 2 eq.) in a mixture of dioxane (12 mL) and water (2 mL) was degassed and purged with nitrogen three times, then the mixture was stirred at 90 °C for 10 h under nitrogen atmosphere. The reaction mixture was concentrated to give the crude product. The residue was purified by semi-preparative reverse phase HPLC (column: [Kromasil 250*50mm*10um; mobile phase: [water (0.1% trifluoroacetic acid)-ACN]; B%: 20ACN% to 50ACN%, 20min). The compound tert-butyl-4-[3-[[4-[3-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl]-2-pyridyl]oxy]cyclobutoxy]piperidine-1-carboxylate (500mg, 0.71mmol, 54% yield, 91% purity) was obtained as a yellow solid.
工程13

ジクロロメタン(5mL)中のtert-ブチル4-[3-[[4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]-2-ピリジル]オキシ]シクロブトキシ]ピペリジン-1-カルボキシレート(500mg、0.78mmol、1当量)の溶液に、塩酸/ジオキサン(4M、5mL、25.75当量)を添加した。混合物を20℃で2時間撹拌した。混合物を濃縮して、粗生成物を得た。化合物2-[6-アミノ-5-[8-[2-[3-(4-ピペリジルオキシ)シクロブトキシ]-4-ピリジル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-イル]ピリダジン-3-イル]フェノール(500mg、粗製、三塩酸塩)を黄色の固体として得た。 To a solution of tert-butyl 4-[3-[[4-[3-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl]-2-pyridyl]oxy]cyclobutoxy]piperidine-1-carboxylate (500 mg, 0.78 mmol, 1 equiv.) in dichloromethane (5 mL) was added hydrochloric acid/dioxane (4 M, 5 mL, 25.75 equiv.). The mixture was stirred at 20°C for 2 h. The mixture was concentrated to give the crude product. The compound 2-[6-amino-5-[8-[2-[3-(4-piperidyloxy)cyclobutoxy]-4-pyridyl]-3,8-diazabicyclo[3.2.1]octan-3-yl]pyridazin-3-yl]phenol (500 mg, crude, trihydrochloride salt) was obtained as a yellow solid.
工程14

ジクロロメタン(16mL)およびメタノール(16mL)の混合物中の2-[6-アミノ-5-[8-[2-[3-(4-ピペリジルオキシ)シクロブトキシ]-4-ピリジル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-イル]ピリダジン-3-イル]フェノール三塩酸塩(400mg、612umol、1当量)および(2S,4R)-4-ヒドロキシ-1-[(2R)-3-メチル-2-[3-(2-オキソエトキシ)イソキサゾール-5-イル]ブタノイル]-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(331mg、0.61mmol、1当量)の溶液に、酢酸ナトリウム(201mg、2.45mmol、4当量)を添加し、次いで、pH=6まで酢酸を添加し、次いで、シアノ水素化ホウ素(192mg、3.06mmol、5当量)を添加した。混合物を20℃で10時間撹拌した。反応混合物を水(20mL)で希釈し、ジクロロメタン(20mL)で3回抽出した。合わせた有機層を無水硫酸ナトリウムで乾燥させ、濃縮して、粗生成物を得、これを半分取逆相HPLC(カラム:Phenomenex Synergi C18 150*25*10um;移動相:[水(0.225%FA)-ACN]B%:18%~48%、9分)によって精製した。(2S,4R)-1-[(2R)-2-[3-[2-[4-[3-[[4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]-2-ピリジル]オキシ]シクロブトキシ]-1-ピペリジル]エトキシ]イソキサゾール-5-イル]-3-メチル-ブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミドビストリフルオロ酢酸塩(321mg、0.28mmol、収率45%)を灰色がかった白色の固体として得た。 2-[6-amino-5-[8-[2-[3-(4-piperidyloxy)cyclobutoxy]-4-pyridyl]-3,8-diazabicyclo[3.2.1]octan-3-yl]pyridazin-3-yl]phenol trihydrochloride (400 mg, 612 umol, 1 equiv.) and (2S,4R)-4-hydroxy-1-[(2R)-3-methyl-2-[3-(2-oxo) To a solution of N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (331 mg, 0.61 mmol, 1 eq.) was added sodium acetate (201 mg, 2.45 mmol, 4 eq.), followed by acetic acid until pH=6, followed by cyanoborohydride (192 mg, 3.06 mmol, 5 eq.). The mixture was stirred at 20° C. for 10 h. The reaction mixture was diluted with water (20 mL) and extracted three times with dichloromethane (20 mL). The combined organic layers were dried over anhydrous sodium sulfate and concentrated to give the crude product, which was purified by semi-preparative reverse phase HPLC (Column: Phenomenex Synergi C18 150*25*10um; Mobile phase: [Water (0.225% FA)-ACN] B%: 18%-48%, 9 min). (2S,4R)-1-[(2R)-2-[3-[2-[4-[3-[[4-[3-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl]-2-pyridyl]oxy]cyclobutoxy]-1-piperidyl]ethoxy]isoxazol-5-yl]-3-methyl-butanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide bistrifluoroacetate (321 mg, 0.28 mmol, 45% yield) was obtained as an off-white solid.
例示的な化合物63および146を、類似の手順を使用して調製した。 Exemplary compounds 63 and 146 were prepared using similar procedures.
例示的な化合物67および68の例示的な合成

工程1

水(500mL)中のNaOH(10g、0.250mol、2.7当量)の撹拌溶液に、ベンジルアルコール(10g、92.47mmol、1当量)、1,4-ジブロモブタン(39.93g、184.9mmol、2.00当量)、およびBu4NHSO4(0.78g、2.297mmol、0.02当量)を室温で添加した。得られた混合物を70℃で2時間撹拌し、次いで、ヘキサン(100mL)で希釈した。得られた混合物を3×50mLの水で洗浄した。有機層を無水硫酸ナトリウムで乾燥させ、減圧下で濃縮した。残渣を、PE/EtOAc(10:1)で溶出するシリカゲルカラムクロマトグラフィーによって精製した。これにより、[(4-ブロモブトキシ)メチル]ベンゼン(10g、44%)を無色の油として得た。 To a stirred solution of NaOH (10 g, 0.250 mol, 2.7 equiv) in water (500 mL), benzyl alcohol (10 g, 92.47 mmol, 1 equiv), 1,4-dibromobutane (39.93 g, 184.9 mmol, 2.00 equiv), and Bu4NHSO4 (0.78 g, 2.297 mmol, 0.02 equiv) were added at room temperature. The resulting mixture was stirred at 70°C for 2 h and then diluted with hexane (100 mL). The resulting mixture was washed with 3 x 50 mL of water. The organic layer was dried over anhydrous sodium sulfate and concentrated under reduced pressure. The residue was purified by silica gel column chromatography eluting with PE/EtOAc (10:1). This gave [(4-bromobutoxy)methyl]benzene (10 g, 44%) as a colorless oil.
工程2

DMF(100mL)中の2-(2-ヒドロキシエトキシ)エタン-1-オール(14.18g、133.7mmol、5当量)の溶液に、水素化ナトリウム(油中60%、2.15g、2.5当量)を窒素雰囲気下で、0℃で添加した。15分後、反応混合物に[(4-ブロモブトキシ)メチル]ベンゼン(6.5g、26.7mmol、1当量)を添加した。得られた混合物を25℃で3時間撹拌し、次いで、水(250mL)を添加することによってクエンチした。得られた混合物をEA(3×200mL)で抽出した。合わせた有機相をブラインで洗浄し、硫酸ナトリウムで乾燥させ、減圧下で濃縮した。残渣を、PE/EtOAc(60%)で溶出するシリカゲルカラムクロマトグラフィーによって精製した。これにより、2-[2-[4-(ベンジルオキシ)ブトキシ]エトキシ]エタン-1-オール(4g、72%)を無色の油として得た。 To a solution of 2-(2-hydroxyethoxy)ethan-1-ol (14.18 g, 133.7 mmol, 5 eq.) in DMF (100 mL) was added sodium hydride (60% in oil, 2.15 g, 2.5 eq.) under nitrogen atmosphere at 0° C. After 15 min, [(4-bromobutoxy)methyl]benzene (6.5 g, 26.7 mmol, 1 eq.) was added to the reaction mixture. The resulting mixture was stirred at 25° C. for 3 h and then quenched by adding water (250 mL). The resulting mixture was extracted with EA (3×200 mL). The combined organic phases were washed with brine, dried over sodium sulfate, and concentrated under reduced pressure. The residue was purified by silica gel column chromatography eluting with PE/EtOAc (60%). This gave 2-[2-[4-(benzyloxy)butoxy]ethoxy]ethan-1-ol (4 g, 72%) as a colorless oil.
工程3
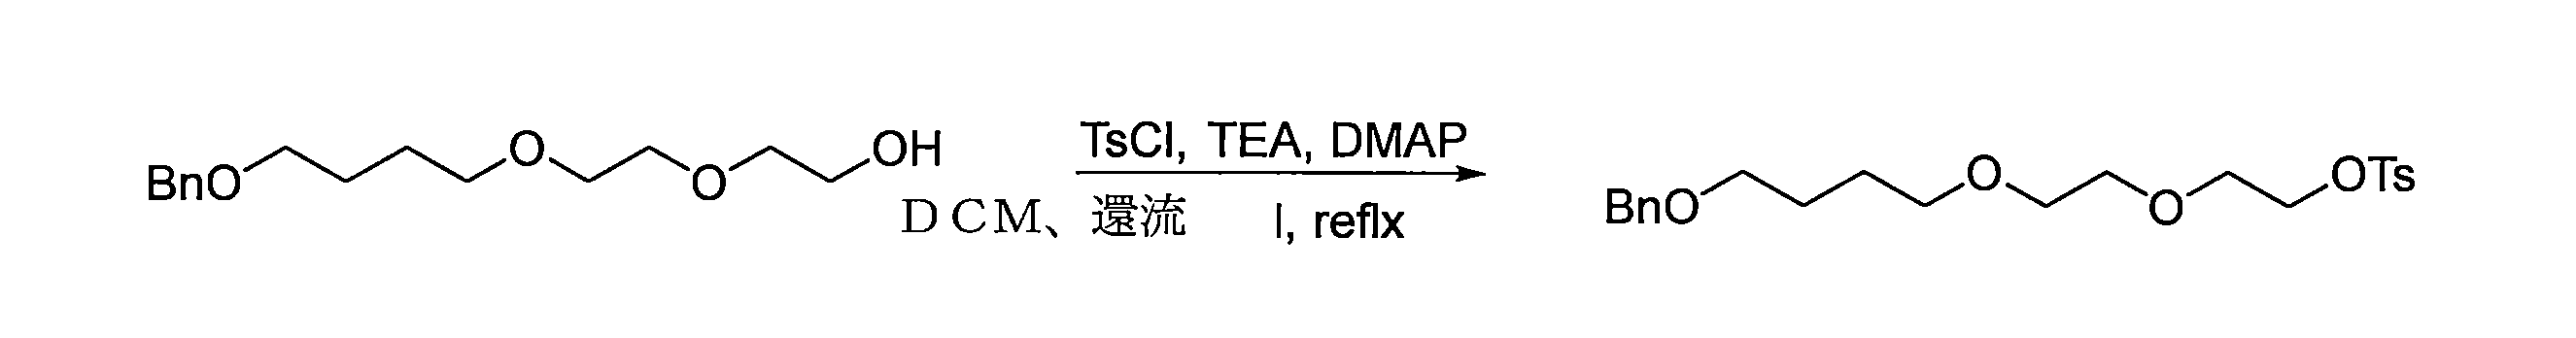
DCM溶液(50mL)中の2-[2-[4-(ベンジルオキシ)ブトキシ]エトキシ]エタン-1-オール(2.4g、8.94mmol、1当量)およびDMAP(218mg、1.79mmol、0.2当量)の撹拌溶液に、TsCl(5.1g、26.83mmol、3当量)およびTEA(2.7g、26.83mmol、3当量)を室温で添加した。得られた混合物を40℃で一晩撹拌した。反応混合物を、DCM(100mL)で希釈し、2×50mLのブラインで洗浄した。有機層を無水硫酸ナトリウムで乾燥させ、減圧下で濃縮した。残渣を、PE/EtOAc(3:1)で溶出するシリカゲルカラムクロマトグラフィーによって精製した。これにより、2-[2-[4-(ベンジルオキシ)ブトキシ]エトキシ]エチル4-メチルベンゼン-1-スルホネート(2.4g、64%)を淡い黄色の固体として得た。 To a stirred solution of 2-[2-[4-(benzyloxy)butoxy]ethoxy]ethan-1-ol (2.4 g, 8.94 mmol, 1 eq.) and DMAP (218 mg, 1.79 mmol, 0.2 eq.) in DCM solution (50 mL), TsCl (5.1 g, 26.83 mmol, 3 eq.) and TEA (2.7 g, 26.83 mmol, 3 eq.) were added at room temperature. The resulting mixture was stirred at 40° C. overnight. The reaction mixture was diluted with DCM (100 mL) and washed with 2×50 mL of brine. The organic layer was dried over anhydrous sodium sulfate and concentrated under reduced pressure. The residue was purified by silica gel column chromatography eluting with PE/EtOAc (3:1). This gave 2-[2-[4-(benzyloxy)butoxy]ethoxy]ethyl 4-methylbenzene-1-sulfonate (2.4 g, 64%) as a pale yellow solid.
工程4

100mLのEtOH中の2-[2-[4-(ベンジルオキシ)ブトキシ]エトキシ]エチル4-メチルベンゼン-1-スルホネート(3.5g、8.28mmol、1当量)の溶液に、10%Pd/C(3.5g)およびCH3COOH(0.95mL、15.88mmol、2.01当量)を窒素雰囲気下で添加した。フラスコを真空化し、水素で流した。得られた混合物を、水素バルーンを使用して水素雰囲気下で、40℃で一晩水素化し、次いで、セライトパッドを通して濾過し、減圧下で濃縮した。残渣を、以下の条件下で逆相カラムクロマトグラフィーによって精製した:カラム、C18シリカゲル;移動相、水中のACN、30分間で10%~50%の勾配。これにより、4-(2-[2-[(4-メチルベンゼンスルホニル)オキシ]エトキシ]エトキシ)ブタン-1-オール(2.2g、80%)を無色の油として得た。 To a solution of 2-[2-[4-(benzyloxy)butoxy]ethoxy]ethyl 4-methylbenzene-1-sulfonate (3.5 g, 8.28 mmol, 1 eq.) in 100 mL of EtOH, 10% Pd/C (3.5 g) and CH3COOH (0.95 mL, 15.88 mmol, 2.01 eq.) were added under nitrogen atmosphere. The flask was evacuated and flushed with hydrogen. The resulting mixture was hydrogenated overnight at 40° C. under hydrogen atmosphere using a hydrogen balloon, then filtered through a celite pad and concentrated under reduced pressure. The residue was purified by reverse phase column chromatography under the following conditions: column, C18 silica gel; mobile phase, ACN in water, 10% to 50% gradient in 30 min. This gave 4-(2-[2-[(4-methylbenzenesulfonyl)oxy]ethoxy]ethoxy)butan-1-ol (2.2 g, 80%) as a colorless oil.
工程5

アセトン(50mL)中のメチル2-(3-ヒドロキシ-1,2-オキサゾール-5-イル)-3-メチルブタノエート(1.5g、7.53mmol、1当量)および4-(2-[2-[(4-メチルベンゼンスルホニル)オキシ]エトキシ]エトキシ)ブタン-1-オール(3.0g、9.04mmol、1.2当量)の撹拌溶液に、Cs2CO3(4.9g、15.04mmol、2.00当量)を、窒素雰囲気下で、室温で添加した。得られた混合物を60℃で4時間撹拌し、次いで、減圧下で濃縮した。残渣を、CH2Cl2/MeOH(10:1)で溶出するシリカゲルカラムクロマトグラフィーによって精製した。これにより、メチル2-(3-[2-[2-(4-ヒドロキシブトキシ)エトキシ]エトキシ]-1,2-オキサゾール-5-イル)-3-メチルブタノエート(800mg、30%)を得た。 To a stirred solution of methyl 2-(3-hydroxy-1,2-oxazol-5-yl)-3-methylbutanoate (1.5 g, 7.53 mmol, 1 equiv.) and 4-(2-[2-[(4-methylbenzenesulfonyl)oxy]ethoxy]ethoxy)butan-1-ol (3.0 g, 9.04 mmol, 1.2 equiv.) in acetone (50 mL), Cs2CO3 (4.9 g, 15.04 mmol, 2.00 equiv.) was added at room temperature under nitrogen atmosphere. The resulting mixture was stirred at 60°C for 4 h and then concentrated under reduced pressure. The residue was purified by silica gel column chromatography eluting with CH2Cl2/MeOH (10:1). This yielded methyl 2-(3-[2-[2-(4-hydroxybutoxy)ethoxy]ethoxy]-1,2-oxazol-5-yl)-3-methylbutanoate (800 mg, 30%).
工程6
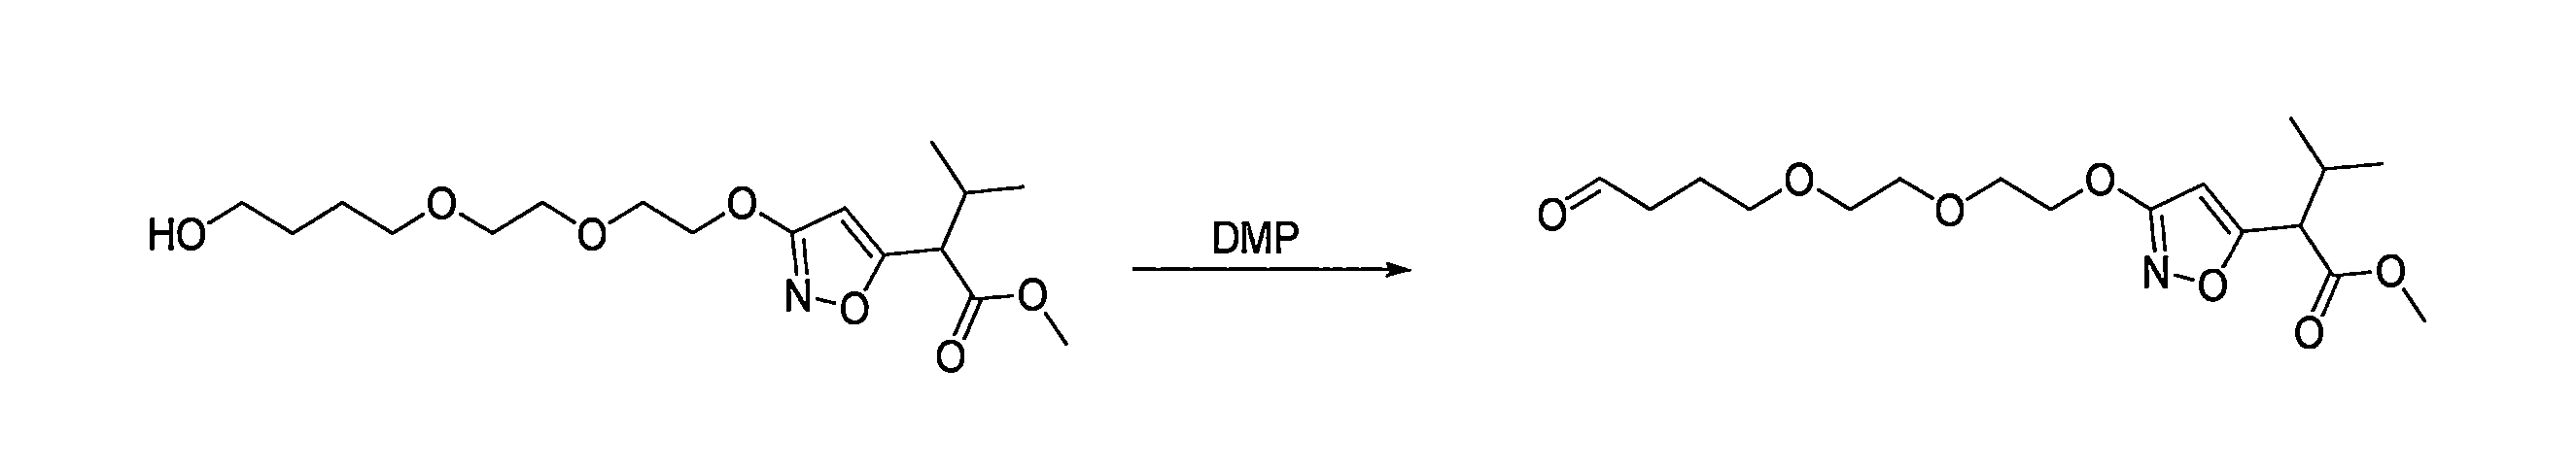
DCM(20mL)中のメチル2-(3-[2-[2-(4-ヒドロキシブトキシ)エトキシ]エトキシ]-1,2-オキサゾール-5-イル)-3-メチルブタノエート(550mg、1.53mmol、1当量)の撹拌溶液に、デスマーチンペルヨージナン(1298mg、3.06mmol、2当量)を0℃で添加した。得られた混合物30℃で2時間撹拌し、次いで、飽和チオ硫酸ナトリウム溶液を添加することによってクエンチした。得られた混合物をDCM(3×20mL)で抽出した。合わせた有機相を飽和重炭酸ナトリウム溶液で洗浄し、硫酸ナトリウムで乾燥させ、減圧下で濃縮した。残渣を、分取TLC(CH2Cl2/MeOH 10:1)によって精製して、メチル3-メチル-2-(3-[2-[2-(4-オキソブトキシ)エトキシ]エトキシ]-1,2-オキサゾール-5-イル)ブタノエート(440mg、80%)を淡い黄色の固体として得た。 To a stirred solution of methyl 2-(3-[2-[2-(4-hydroxybutoxy)ethoxy]ethoxy]-1,2-oxazol-5-yl)-3-methylbutanoate (550 mg, 1.53 mmol, 1 equiv.) in DCM (20 mL) was added Dess-Martin periodinane (1298 mg, 3.06 mmol, 2 equiv.) at 0 °C. The resulting mixture was stirred at 30 °C for 2 h and then quenched by adding saturated sodium thiosulfate solution. The resulting mixture was extracted with DCM (3 × 20 mL). The combined organic phases were washed with saturated sodium bicarbonate solution, dried over sodium sulfate and concentrated under reduced pressure. The residue was purified by preparative TLC (CH2Cl2/MeOH 10:1) to give methyl 3-methyl-2-(3-[2-[2-(4-oxobutoxy)ethoxy]ethoxy]-1,2-oxazol-5-yl)butanoate (440 mg, 80%) as a pale yellow solid.
一連の他の実施例(例えば、例示的な化合物91)について記載されるものと同様の、ならびに当業者に明らかである手順を使用して、以下のスキームに従って、メチル3-メチル-2-(3-[2-[2-(4-オキソブトキシ)エトキシ]エトキシ]-1,2-オキサゾール-5-イル)ブタノエートを最終化合物に変換した。

例示的な化合物65、66、71、73、および74を、類似の手順を使用して調製した。 Exemplary compounds 65, 66, 71, 73, and 74 were prepared using similar procedures.
例示的な化合物69および70の例示的な合成

工程1

250mLの丸底フラスコに、エタノール(100mL)中の3-(ベンジルオキシ)シクロブタン-1-オン(6.0g、34.05mmol、1.0当量)の溶液を入れた。これに続いて、0℃で、水素化ホウ素ナトリウム(1.25g、33.94mmol、1.00当量)を数回に分けて添加した。得られた溶液を、0℃で3時間撹拌した。次いで、反応物を、水(100mL)を添加することによってクエンチした。得られた溶液を、酢酸エチル(50mL×2)で抽出し、有機層を合わせ、無水硫酸ナトリウムで乾燥させ、真空下で濃縮した。これにより、6.0g(99%)の(1s,3s)-3-(ベンジルオキシ)シクロブタン-1-オールを淡い黄色の油として得た。 A 250 mL round bottom flask was charged with a solution of 3-(benzyloxy)cyclobutan-1-one (6.0 g, 34.05 mmol, 1.0 equiv) in ethanol (100 mL). This was followed by the addition of sodium borohydride (1.25 g, 33.94 mmol, 1.00 equiv) in portions at 0° C. The resulting solution was stirred at 0° C. for 3 hours. The reaction was then quenched by the addition of water (100 mL). The resulting solution was extracted with ethyl acetate (50 mL×2) and the organic layers were combined, dried over anhydrous sodium sulfate, and concentrated under vacuum. This afforded 6.0 g (99%) of (1s,3s)-3-(benzyloxy)cyclobutan-1-ol as a pale yellow oil.
工程2

窒素の不活性雰囲気でパージし、維持した500mLの丸底フラスコに、N,N-ジメチルホルムアミド(100mL)中の(1s,3s)-3-(ベンジルオキシ)シクロブタン-1-オール(8.0g、44.89mmol、1.10当量)の溶液を入れた。続いて、水素化ナトリウム(60%)(2.15g、89.58mmol、1.20当量)を0℃で数回に分けて添加した。これに、4-ブロモ-2-フルオロピリジン(70.0g、39.78mmol、1.00当量)を添加した。得られた溶液を、室温で一晩撹拌した。次いで、反応物を、水/氷を添加することによってクエンチした。得られた溶液を酢酸エチル(80mL×2)で抽出し、有機層を合わせた。得られた混合物をブライン(80mL×2)で洗浄した。結果として生じる混合物を減圧下で濃縮した。残渣を、酢酸エチル/石油エーテル(1:1)とともにシリカゲルカラムに充填させた。これにより、7.0g(53%)の4-ブロモ-2-[(1s,3s)-3-(ベンジルオキシ)シクロブトキシ]ピリジンを淡い黄色の油として得た。 A solution of (1s,3s)-3-(benzyloxy)cyclobutan-1-ol (8.0 g, 44.89 mmol, 1.10 equiv.) in N,N-dimethylformamide (100 mL) was placed in a 500 mL round bottom flask purged and maintained with an inert atmosphere of nitrogen. Sodium hydride (60%) (2.15 g, 89.58 mmol, 1.20 equiv.) was then added in portions at 0°C. To this was added 4-bromo-2-fluoropyridine (70.0 g, 39.78 mmol, 1.00 equiv.). The resulting solution was stirred overnight at room temperature. The reaction was then quenched by adding water/ice. The resulting solution was extracted with ethyl acetate (80 mL x 2) and the organic layers were combined. The resulting mixture was washed with brine (80 mL x 2). The resulting mixture was concentrated under reduced pressure. The residue was loaded onto a silica gel column with ethyl acetate/petroleum ether (1:1). This yielded 7.0 g (53%) of 4-bromo-2-[(1s,3s)-3-(benzyloxy)cyclobutoxy]pyridine as a pale yellow oil.
工程3

窒素の不活性雰囲気でパージし、維持した500mLの丸底フラスコに、トルエン(200mL)中の4-ブロモ-2-[(1s,3s)-3-(ベンジルオキシ)シクロブトキシ]ピリジン(8.4g、25.13mmol、1.30当量)、tert-ブチル3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレート(4.11g、19.36mmol、1.00当量)、tert-ブトキシドナトリウム(3.2g、33.2mmol、1.5当量)、キサントホス(2.32g、4.01mmol、0.20当量)、Pd2(dba)3CH2Cl2(430.0mg、0.40mmol、0.02当量)の溶液を入れた。得られた溶液を100℃で一晩撹拌した。次いで、反応物を、水(100mL)を添加することによってクエンチした。得られた溶液を酢酸エチルで抽出し(80mL×2)、有機層を合わせた。得られた混合物をブライン(100mL×2)で洗浄した。混合物を無水硫酸ナトリウムで乾燥させ、減圧下で濃縮した。残渣を、酢酸エチル/ヘキサン(1:1)で溶出するシリカゲルカラム上に適用した。これにより、8.0g(89%)のtert-ブチル8-[2-[(1s,3s)-3-(ベンジルオキシ)シクロブトキシ]ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレートを淡い黄色の油として得た。 A 500 mL round bottom flask purged and maintained with an inert atmosphere of nitrogen was charged with a solution of 4-bromo-2-[(1s,3s)-3-(benzyloxy)cyclobutoxy]pyridine (8.4 g, 25.13 mmol, 1.30 equiv.), tert-butyl 3,8-diazabicyclo[3.2.1]octane-3-carboxylate (4.11 g, 19.36 mmol, 1.00 equiv.), sodium tert-butoxide (3.2 g, 33.2 mmol, 1.5 equiv.), Xantphos (2.32 g, 4.01 mmol, 0.20 equiv.), Pd2(dba)3CH2Cl2 (430.0 mg, 0.40 mmol, 0.02 equiv.) in toluene (200 mL). The resulting solution was stirred at 100 °C overnight. The reaction was then quenched by adding water (100 mL). The resulting solution was extracted with ethyl acetate (80 mL x 2) and the organic layers were combined. The resulting mixture was washed with brine (100 mL x 2). The mixture was dried over anhydrous sodium sulfate and concentrated under reduced pressure. The residue was applied onto a silica gel column eluted with ethyl acetate/hexane (1:1). This gave 8.0 g (89%) of tert-butyl 8-[2-[(1s,3s)-3-(benzyloxy)cyclobutoxy]pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octane-3-carboxylate as a pale yellow oil.
工程4

150mLのエタノール中のtert-ブチル8-[2-[(1s,3s)-3-(ベンジルオキシ)シクロブトキシ]ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレート(7.0g、15.03mmol、1.00当量)の溶液に、窒素雰囲気下で、250mLの丸底フラスコ中で水酸化パラジウム触媒(7.0g、49.85mmol、3.32当量)および酢酸(2mL)を添加した。次いで、フラスコを真空化し、水素でフラッシュした。反応混合物を、水素バルーンを使用して水素雰囲気下で、50℃で3日間水素化し、次いで、セライトパッドを通して濾過し、減圧下で濃縮した。これにより、5.4g(96%)のtert-ブチル8-[2-[(1s,3s)-3-ヒドロキシシクロブトキシ]ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレートを淡い黄色の脂として得た。 To a solution of tert-butyl 8-[2-[(1s,3s)-3-(benzyloxy)cyclobutoxy]pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octane-3-carboxylate (7.0 g, 15.03 mmol, 1.00 equiv.) in 150 mL of ethanol was added palladium hydroxide catalyst (7.0 g, 49.85 mmol, 3.32 equiv.) and acetic acid (2 mL) in a 250 mL round-bottom flask under nitrogen atmosphere. The flask was then evacuated and flushed with hydrogen. The reaction mixture was hydrogenated under hydrogen atmosphere at 50° C. for 3 days using a hydrogen balloon, then filtered through a celite pad and concentrated under reduced pressure. This yielded 5.4 g (96%) of tert-butyl 8-[2-[(1s,3s)-3-hydroxycyclobutoxy]pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octane-3-carboxylate as a pale yellow oil.
工程5
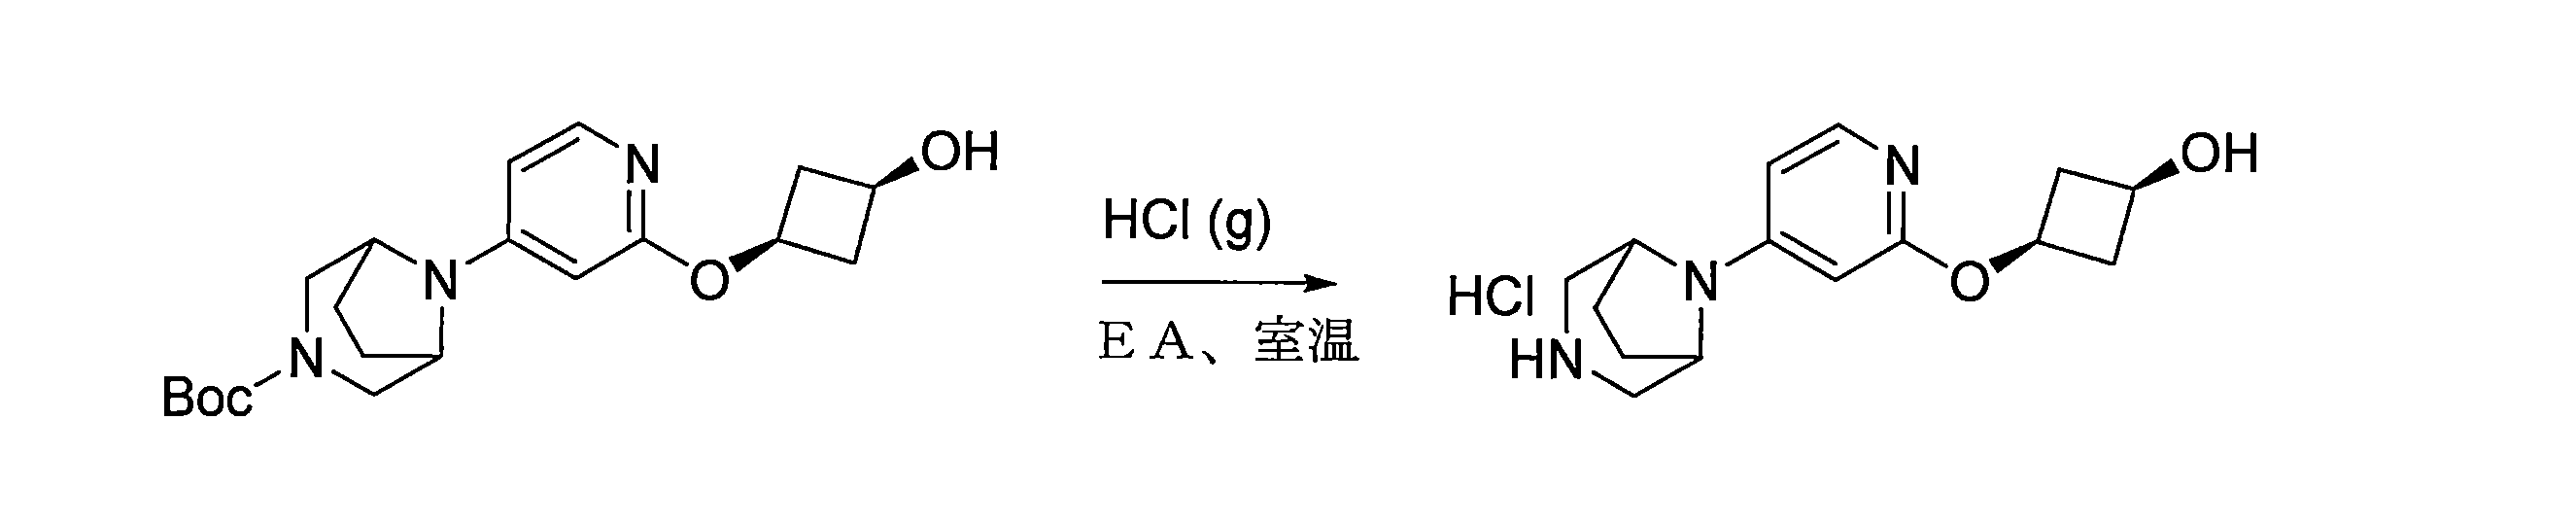
100mLの丸底フラスコに、酢酸エチルおよびメタノール(10:1)(50mL)の混合溶媒中のtert-ブチル8-[2-[(1s,3s)-3-ヒドロキシシクロブトキシ]ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレート(5.0g、13.32mmol、1.00当量)の溶液を入れた。塩化水素(g)を通過させた。得られた混合物を室温で3時間撹拌し、減圧下で濃縮した。これにより、4.0g(96%)の(1s,3s)-3-[(4-[3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)オキシ]シクロブタン-1-オール塩酸塩を灰色がかった白色の固体として得た。 A 100 mL round bottom flask was charged with a solution of tert-butyl 8-[2-[(1s,3s)-3-hydroxycyclobutoxy]pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octane-3-carboxylate (5.0 g, 13.32 mmol, 1.00 equiv.) in a mixture of ethyl acetate and methanol (10:1) (50 mL). Hydrogen chloride (g) was passed through. The resulting mixture was stirred at room temperature for 3 h and concentrated under reduced pressure. This gave 4.0 g (96%) of (1s,3s)-3-[(4-[3,8-diazabicyclo[3.2.1]octane-8-yl]pyridin-2-yl)oxy]cyclobutan-1-ol hydrochloride as an off-white solid.
工程6
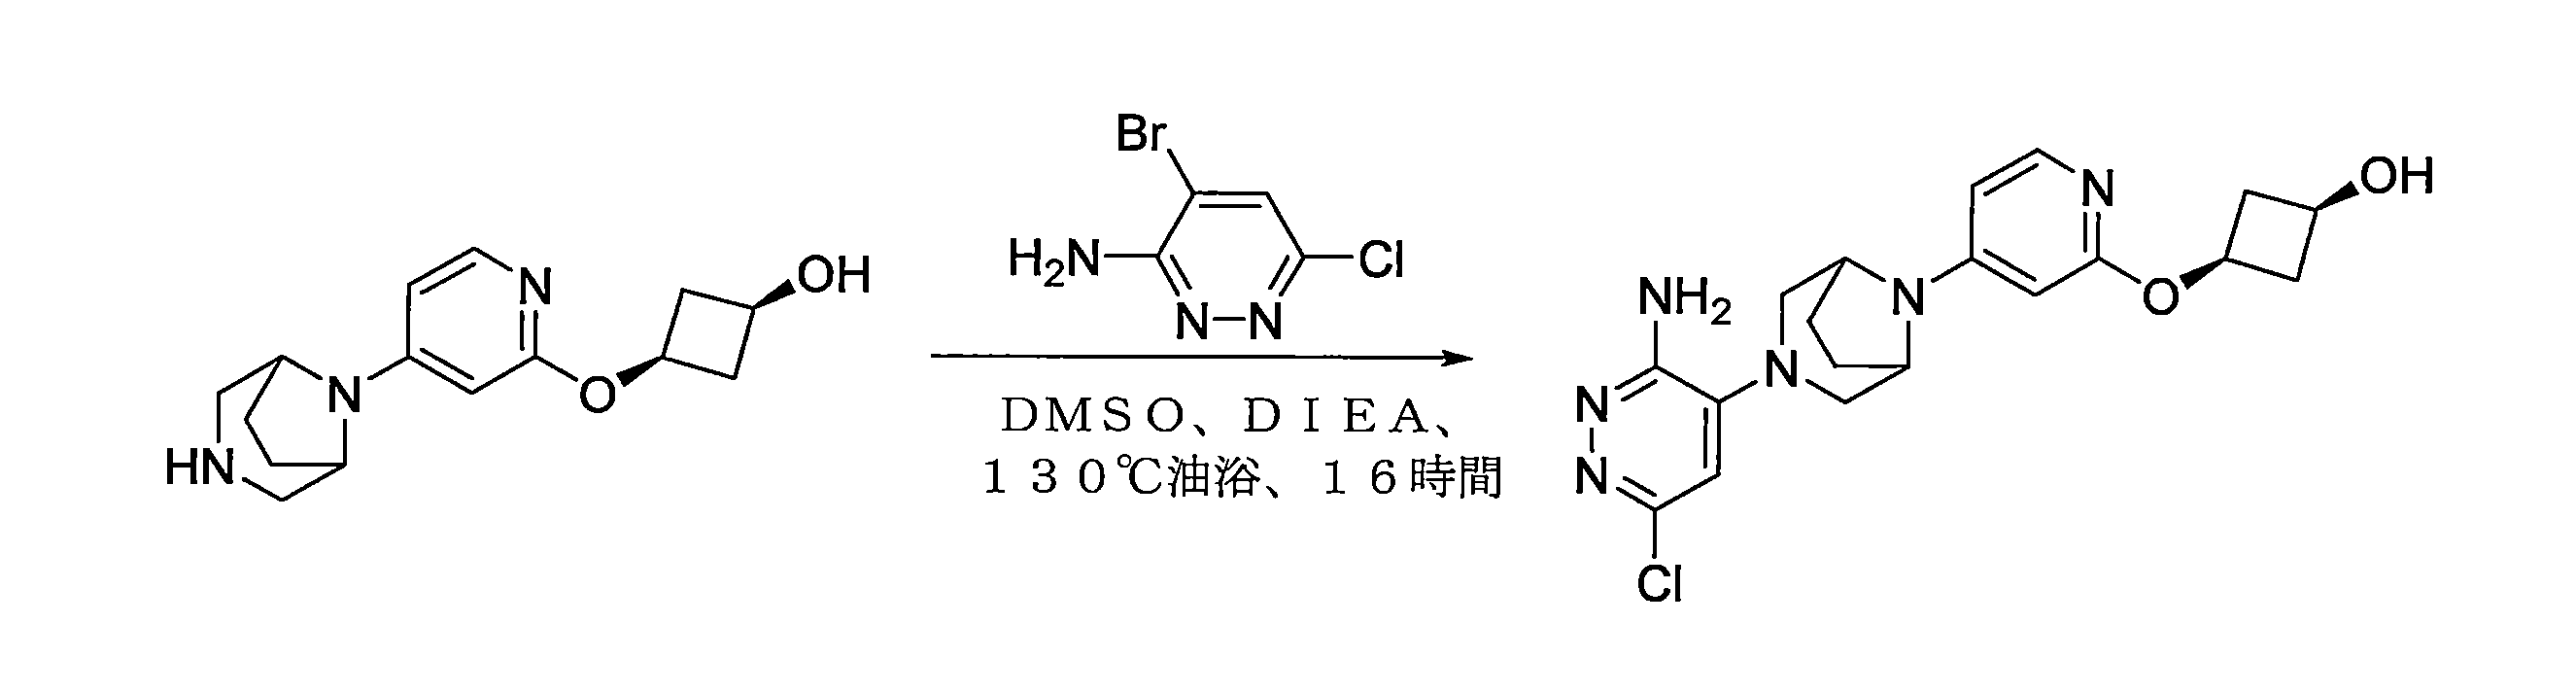
窒素の不活性雰囲気でパージし、維持した20mLの圧力タンク反応器に、ジメチルスルホキシド(10mL)中の(1s,3s)-3-[(4-[3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)オキシ]シクロブタン-1-オール塩酸塩(1.0g、3.21mmol、1.00当量)の溶液を入れた。次いで、N,N-ジイソプロピルエチルアミン(5mL)および4-ブロモ-6-クロロピリダジン-3-アミン(3.01g、14.44mmol、4.00当量)を添加した。得られた溶液を130℃で16時間撹拌した。次いで、反応物を、水(50mL)を添加することによってクエンチした。得られた溶液を酢酸エチル(3×30mL)で抽出し、有機層を合わせた。得られた混合物をブライン(3×50mL)で洗浄した。混合物を、無水硫酸ナトリウムで乾燥させた。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。これにより、490mg(38%)の(1s,3s)-3-([4-[3-(3-アミノ-6-クロロピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル]オキシ)シクロブタン-1-オールを黄色の油として得た。 A 20 mL pressure tank reactor purged and maintained with an inert atmosphere of nitrogen was charged with a solution of (1s,3s)-3-[(4-[3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl)oxy]cyclobutan-1-ol hydrochloride (1.0 g, 3.21 mmol, 1.00 equiv.) in dimethylsulfoxide (10 mL). N,N-diisopropylethylamine (5 mL) and 4-bromo-6-chloropyridazin-3-amine (3.01 g, 14.44 mmol, 4.00 equiv.) were then added. The resulting solution was stirred at 130° C. for 16 hours. The reaction was then quenched by adding water (50 mL). The resulting solution was extracted with ethyl acetate (3×30 mL) and the organic layers were combined. The resulting mixture was washed with brine (3×50 mL). The mixture was dried over anhydrous sodium sulfate. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (10:1). This gave 490 mg (38%) of (1s,3s)-3-([4-[3-(3-amino-6-chloropyridazin-4-yl)-3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl]oxy)cyclobutan-1-ol as a yellow oil.
工程7

窒素の不活性雰囲気でパージし、維持した20-mLの圧力タンク反応器に、ジオキサンおよび水(4:1)(15mL)、[2-(メトキシメトキシ)フェニル]ボロン酸(543.5mg、2.99mmol、1.50当量)、炭酸カリウム(823.9mg、6.0mmol、3.00当量)、Pd(PPh3)4(230.0mg、0.20mmol、0.10当量)の混合溶媒中の(1s,3s)-3-([4-[3-(3-アミノ-6-クロロピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル]オキシ)シクロブタン-1-オール(800.0mg、1.99mmol、1.00当量)の溶液を入れた。得られた溶液を90℃で2時間撹拌した。次いで、反応物を、水(30mL)を添加することによってクエンチした。得られた溶液を酢酸エチル(50mL×2)で抽出し、有機層を合わせた。得られた混合物をブライン(50mL×2)で洗浄した。混合物を、無水硫酸ナトリウムで乾燥させた。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。これにより、820.0mg(82%)の(1s,3s)-3-[[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]オキシ]シクロブタン-1-オールを黄色の固体として得た。 A 20-mL pressure tank reactor purged and maintained with an inert atmosphere of nitrogen was charged with a solution of (1s,3s)-3-([4-[3-(3-amino-6-chloropyridazin-4-yl)-3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl]oxy)cyclobutan-1-ol (800.0 mg, 1.99 mmol, 1.00 equiv.) in a mixture of dioxane and water (4:1) (15 mL), [2-(methoxymethoxy)phenyl]boronic acid (543.5 mg, 2.99 mmol, 1.50 equiv.), potassium carbonate (823.9 mg, 6.0 mmol, 3.00 equiv.), and Pd(PPh3)4 (230.0 mg, 0.20 mmol, 0.10 equiv.). The resulting solution was stirred at 90 °C for 2 h. The reaction was then quenched by adding water (30 mL). The resulting solution was extracted with ethyl acetate (50 mL x 2) and the organic layers were combined. The resulting mixture was washed with brine (50 mL x 2). The mixture was dried over anhydrous sodium sulfate. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (10:1). This gave 820.0 mg (82%) of (1s,3s)-3-[[4-(3-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl]oxy]cyclobutan-1-ol as a yellow solid.
工程8
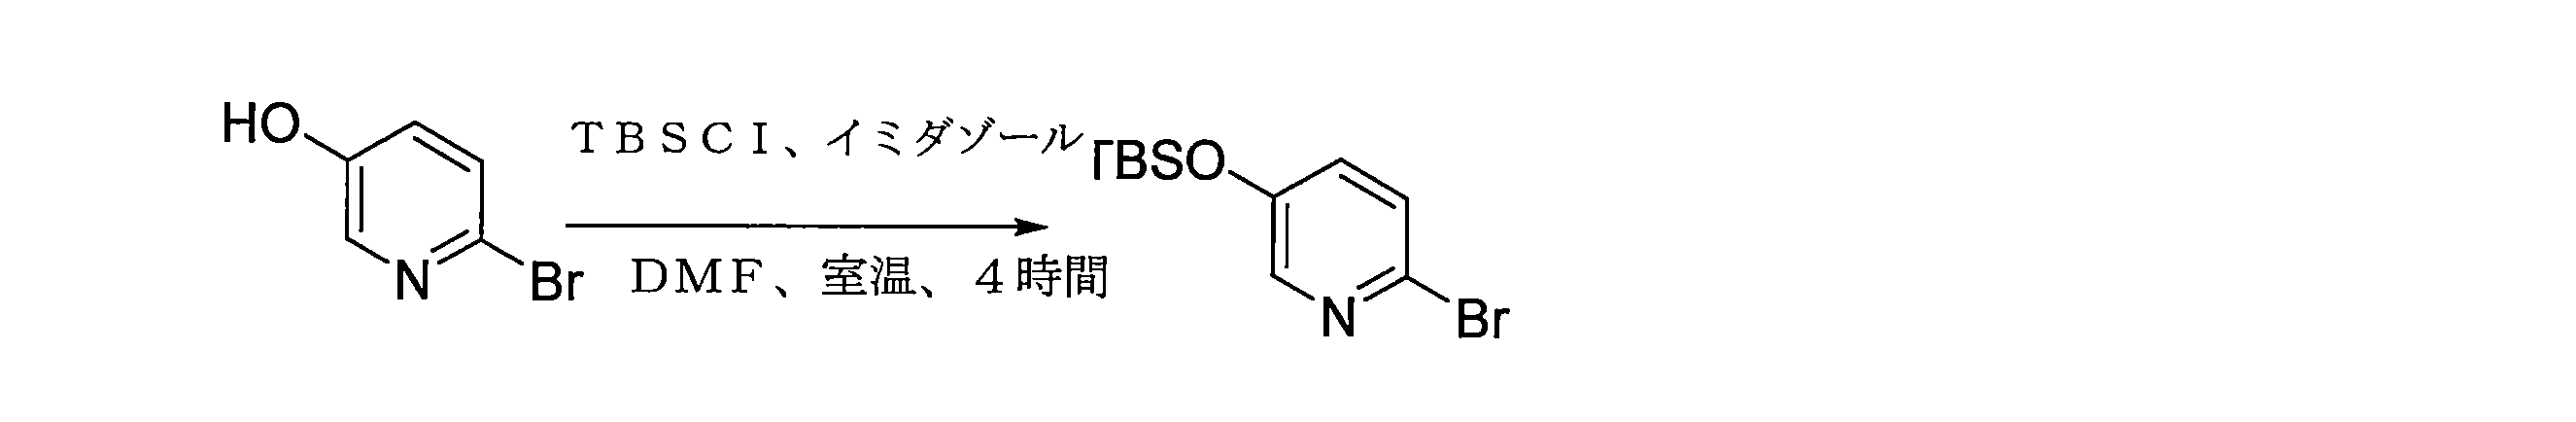
DMF(100mL)中の6-ブロモピリジン-3-オール(12.70g、72.9mmol、1.00当量)およびイミダゾール(7.45g、109.5mmol、1.50当量)の撹拌溶液に、TBSCl(16.50g、109.47mmol、1.50当量)を窒素雰囲気下で、0℃で少しずつ添加した。得られた混合物を、窒素雰囲気下、室温で3時間撹拌した。反応物を、室温で、水(100mL)でクエンチし、得られた混合物をEtOAc(50mL×3)で抽出した。合わせた有機層をブライン(100mL×2)で洗浄し、無水硫酸ナトリウムで乾燥させ、減圧下で濃縮した。残渣を、PE/EtOAc(5:1)で溶出するシリカゲルカラムクロマトグラフィーによって精製し、2-ブロモ-5-[(tert-ブチルジメチルシリル)オキシ]ピリジン(19g、90%)を無色の油として得た。 To a stirred solution of 6-bromopyridin-3-ol (12.70 g, 72.9 mmol, 1.00 equiv.) and imidazole (7.45 g, 109.5 mmol, 1.50 equiv.) in DMF (100 mL), TBSCl (16.50 g, 109.47 mmol, 1.50 equiv.) was added portionwise under nitrogen atmosphere at 0°C. The resulting mixture was stirred at room temperature under nitrogen atmosphere for 3 h. The reaction was quenched with water (100 mL) at room temperature and the resulting mixture was extracted with EtOAc (50 mL x 3). The combined organic layers were washed with brine (100 mL x 2), dried over anhydrous sodium sulfate and concentrated under reduced pressure. The residue was purified by silica gel column chromatography eluting with PE/EtOAc (5:1) to give 2-bromo-5-[(tert-butyldimethylsilyl)oxy]pyridine (19 g, 90%) as a colorless oil.
工程9

1,4-ジオキサン(20mL)中の2-ブロモ-5-[(tert-ブチルジメチルシリル)オキシ]ピリジン(4.00g、13.88mmol、1.00当量)およびNaI(10.40g、69.38mmol、5.00当量)の撹拌混合物に、CuI(0.26g、1.365mmol、0.10当量)およびメチル[2-(メチルアミノ)エチル]アミン(0.12g、1.361mmol、0.10当量)を窒素雰囲気下で、室温で、滴下で添加した。得られた混合物を窒素雰囲気下で、110℃で5時間撹拌した。反応混合物を濾過し、フィルターケーキを1,4-ジオキサンですすいだ。濾液を減圧下で濃縮させた。残渣を、ヘキサン/EtOAc(5:1)で溶出するシリカゲルカラムクロマトグラフィーによって精製して、5-[(tert-ブチルジメチルシリル)オキシ]-2-ヨードピリジン(4.2g、90%)を灰色がかった白色の固体として得た。 To a stirred mixture of 2-bromo-5-[(tert-butyldimethylsilyl)oxy]pyridine (4.00 g, 13.88 mmol, 1.00 equiv.) and NaI (10.40 g, 69.38 mmol, 5.00 equiv.) in 1,4-dioxane (20 mL), CuI (0.26 g, 1.365 mmol, 0.10 equiv.) and methyl[2-(methylamino)ethyl]amine (0.12 g, 1.361 mmol, 0.10 equiv.) were added dropwise at room temperature under a nitrogen atmosphere. The resulting mixture was stirred at 110° C. for 5 h under a nitrogen atmosphere. The reaction mixture was filtered and the filter cake was rinsed with 1,4-dioxane. The filtrate was concentrated under reduced pressure. The residue was purified by silica gel column chromatography eluting with hexane/EtOAc (5:1) to give 5-[(tert-butyldimethylsilyl)oxy]-2-iodopyridine (4.2 g, 90%) as an off-white solid.
工程10

THF(20mL)中の5-[(tert-ブチルジメチルシリル)オキシ]-2-ヨードピリジン(4.20g、12.53mmol、1.00当量)の撹拌溶液に、TBAF(1mol/L、27mL、27mmol、2.00当量)を水素雰囲気下で、0℃で、滴下で添加した。得られた混合物を窒素雰囲気下で、室温で2時間撹拌した。反応物を室温で、水(50mL)でクエンチした。得られた混合物をEtOAc(50mL×3)で抽出した。合わせた有機相をブライン(50mL)で洗浄し、硫酸ナトリウムで乾燥させ、減圧下で濃縮した。残渣を、ヘキサン/EtOAc(2:1)で溶出するシリカゲルカラムクロマトグラフィーによって精製して、6-ヨードピリジン-3-オール(2.3g、83%)を淡い黄色の固体として得た。 To a stirred solution of 5-[(tert-butyldimethylsilyl)oxy]-2-iodopyridine (4.20 g, 12.53 mmol, 1.00 equiv.) in THF (20 mL) was added TBAF (1 mol/L, 27 mL, 27 mmol, 2.00 equiv.) dropwise under hydrogen atmosphere at 0°C. The resulting mixture was stirred at room temperature under nitrogen atmosphere for 2 h. The reaction was quenched with water (50 mL) at room temperature. The resulting mixture was extracted with EtOAc (50 mL x 3). The combined organic phase was washed with brine (50 mL), dried over sodium sulfate, and concentrated under reduced pressure. The residue was purified by silica gel column chromatography eluting with hexane/EtOAc (2:1) to give 6-iodopyridin-3-ol (2.3 g, 83%) as a pale yellow solid.
工程11

THF(10mL)中のPPh3(523mg、2.00mmol、2.00当量)の溶液に、DIAD(403mg、2.00mmol、2.00当量)、(1s,3s)-3-[[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]オキシ]シクロブタン-1-オール(504mg、1.00mmol、1当量)、および6-ヨードピリジン-3-オール(265mg、1.20mmol、1.20当量)を窒素雰囲気下で、0℃で添加した。得られた混合物を0℃で5分間撹拌し、50℃まで加熱し、窒素雰囲気下で、50℃で2時間撹拌した。反応物を、室温で水(20mL)を添加することによってクエンチし、得られた混合物をEtOAc(20mL×3)で抽出した。合わせた有機相をブライン(30mL)で洗浄し、硫酸ナトリウムで乾燥させ、減圧下で濃縮した。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラムクロマトグラフィーによって精製して、6-[2-(メトキシメトキシ)フェニル]-4-(8-[2-[(1r,3r)-3-[(6-ヨードピリジン-3-イル)オキシ]シクロブトキシ]ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-イル)ピリダジン-3-アミン(382mg、54%)を淡い黄色の固体として得た。 To a solution of PPh3 (523 mg, 2.00 mmol, 2.00 equiv.) in THF (10 mL), DIAD (403 mg, 2.00 mmol, 2.00 equiv.), (1s,3s)-3-[[4-(3-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl]oxy]cyclobutan-1-ol (504 mg, 1.00 mmol, 1 equiv.), and 6-iodopyridin-3-ol (265 mg, 1.20 mmol, 1.20 equiv.) were added at 0°C under nitrogen atmosphere. The resulting mixture was stirred at 0°C for 5 min, heated to 50°C, and stirred at 50°C for 2 h under nitrogen atmosphere. The reaction was quenched by adding water (20 mL) at room temperature, and the resulting mixture was extracted with EtOAc (20 mL x 3). The combined organic phase was washed with brine (30 mL), dried over sodium sulfate, and concentrated under reduced pressure. The residue was purified by silica gel column chromatography eluting with dichloromethane/methanol (10:1) to give 6-[2-(methoxymethoxy)phenyl]-4-(8-[2-[(1r,3r)-3-[(6-iodopyridin-3-yl)oxy]cyclobutoxy]pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octan-3-yl)pyridazin-3-amine (382 mg, 54%) as a pale yellow solid.
工程12

アセトン(40mL)中の(2S,4R)-4-ヒドロキシ-1-[2-(3-ヒドロキシ-1,2-オキサゾール-5-イル)-3-メチルブタノイル]-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(780mg、1.56mmol、1.00当量)の溶液に、Cs2CO3(1019mg、3.13mmol、2.00当量)、および3-ブロモプロパ-1-イン(558mg、4.69mmol、3.00当量)を室温で添加した。得られた混合物を窒素雰囲気下で、50℃で3時間撹拌した。混合物を濾過し、フィルターケーキをTHF(50mL×2)ですすいだ。濾液を減圧下で濃縮させた。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラムクロマトグラフィーによって精製して、(2S,4R)-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]-1-[3-メチル-2-[3-(プロパ-2-イン-1-イルオキシ)-1,2-オキサゾール-5-イル]ブタノイル]ピロリジン-2-カルボキサミド(520mg、62%)を灰色がかった白色の固体として得た。 To a solution of (2S,4R)-4-hydroxy-1-[2-(3-hydroxy-1,2-oxazol-5-yl)-3-methylbutanoyl]-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (780 mg, 1.56 mmol, 1.00 equiv.) in acetone (40 mL), Cs2CO3 (1019 mg, 3.13 mmol, 2.00 equiv.), and 3-bromoprop-1-yne (558 mg, 4.69 mmol, 3.00 equiv.) were added at room temperature. The resulting mixture was stirred at 50°C for 3 h under a nitrogen atmosphere. The mixture was filtered and the filter cake was rinsed with THF (50 mL x 2). The filtrate was concentrated under reduced pressure. The residue was purified by silica gel column chromatography eluting with dichloromethane/methanol (10:1) to give (2S,4R)-4-hydroxy-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]-1-[3-methyl-2-[3-(prop-2-yn-1-yloxy)-1,2-oxazol-5-yl]butanoyl]pyrrolidine-2-carboxamide (520 mg, 62%) as an off-white solid.
工程13

DMF(5mL)中の6-[2-(メトキシメトキシ)フェニル]-4-(8-[2-[(1r,3r)-3-[(6-ヨードピリジン-3-イル)オキシ]シクロブトキシ]ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-イル)ピリダジン-3-アミン(224mg、0.317mmol、1.00当量)およびCuI(6.0mg、0.032mmol、0.10当量)の撹拌混合物に、TEA(96mg、0.950mmol、3.00当量)、およびPd(PPh3)2Cl2(22.2mg、0.032mmol、0.10当量)を窒素雰囲気下で、室温で添加した。得られた混合物を室温で5分間撹拌し、次いで、DMF(2mL)中の(2S,4R)-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]-1-[3-メチル-2-[3-(プロパ-2-イン-1イルオキシ)-1,2-オキサゾール-5-イル]ブタノイル]ピロリジン-2-カルボキサミド(170mg、0.317mmol、1.00当量)の溶液に添加した。反応混合物を窒素雰囲気下で、65℃で2時間撹拌した。反応物を、水(30mL)を添加することによってクエンチした。得られた混合物をEtOAc(20mL×3)で抽出した。合わせた有機層をブライン(30mL×3)で洗浄し、無水硫酸ナトリウムで乾燥させ、減圧下で濃縮した。残渣を、ジクロロメタン/メタノール(9:1)で溶出する分取TLCによって精製して、(2S,4R)-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]-1-(3-メチル-2-[3-[(3-[5-[(1r,3r)-3-[[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]オキシ]シクロブトキシ]ピリジン-2-イル]プロパ-2-イン-1-イル)オキシ]-1,2-オキサゾール-5-イル]ブタノイル)ピロリジン-2-カルボキサミド(175mg、49%)を淡い黄色の固体として得た。 To a stirred mixture of 6-[2-(methoxymethoxy)phenyl]-4-(8-[2-[(1r,3r)-3-[(6-iodopyridin-3-yl)oxy]cyclobutoxy]pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octan-3-yl)pyridazin-3-amine (224 mg, 0.317 mmol, 1.00 equiv.) and CuI (6.0 mg, 0.032 mmol, 0.10 equiv.) in DMF (5 mL) was added TEA (96 mg, 0.950 mmol, 3.00 equiv.) and Pd(PPh3)2Cl2 (22.2 mg, 0.032 mmol, 0.10 equiv.) under a nitrogen atmosphere at room temperature. The resulting mixture was stirred at room temperature for 5 minutes and then added to a solution of (2S,4R)-4-hydroxy-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]-1-[3-methyl-2-[3-(prop-2-yn-1yloxy)-1,2-oxazol-5-yl]butanoyl]pyrrolidine-2-carboxamide (170 mg, 0.317 mmol, 1.00 equiv) in DMF (2 mL). The reaction mixture was stirred at 65° C. for 2 hours under nitrogen atmosphere. The reaction was quenched by adding water (30 mL). The resulting mixture was extracted with EtOAc (20 mL×3). The combined organic layers were washed with brine (30 mL×3), dried over anhydrous sodium sulfate and concentrated under reduced pressure. The residue was purified by preparative TLC eluting with dichloromethane/methanol (9:1) to give (2S,4R)-4-hydroxy-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]-1-(3-methyl-2-[3-[(3-[5-[(1r,3r)-3-[[4-(3-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl]oxy]cyclobutoxy]pyridin-2-yl]prop-2-yn-1-yl)oxy]-1,2-oxazol-5-yl]butanoyl)pyrrolidine-2-carboxamide (175 mg, 49%) as a pale yellow solid.
(2S,4R)-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]-1-(3-メチル-2-[3-[(3-[5-[(1r,3r)-3-[(4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)オキシ]シクロブトキシ]ピリジン-2-イル]プロパ-2-イン-1-イル)オキシ]-1,2-オキサゾール-5-イル]ブタノイル)ピロリジン-2-カルボキサミド(620mg)の2つのジアステレオマーを、以下の条件でキラルHPLCによって分離した。カラム:CHIRALPAK IG、2.0cm I.D*25cm(5um);移動相A:MeOH(8mmol/L NH3.MeOH)、移動相B:DCM、流速:20mL/分、勾配:17分間で30%B~30%B;220/254nm;RT1:8.495分;RT2:12.854分。これにより、(2S,4R)-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]-1-[(2S)-3-メチル-2-[3-[(3-[5-[(1r,3r)-3-[(4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)オキシ]シクロブトキシ]ピリジン-2-イル]プロパ-2-イン-1-イル)オキシ]-1,2-オキサゾール-5-イル]ブタノイル]ピロリジン-2-カルボキサミド(淡い黄色の固体)として暫定的に特定された137mg(22%)の異性体1、および(2S,4R)-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]-1-[(2R)-3-メチル-2-[3-[(3-[5-[(1r,3r)-3-[(4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]ピリジン-2-イル)オキシ]シクロブトキシ]ピリジン-2-イル]プロパ-2-イン-1-イル)オキシ]-1,2-オキサゾール-5-イル]ブタノイル]ピロリジン-2-カルボキサミド(灰色がかった白色の固体)として暫定的に特定された199mg(32%)の異性体2を得た。 The two diastereomers of (2S,4R)-4-hydroxy-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]-1-(3-methyl-2-[3-[(3-[5-[(1r,3r)-3-[(4-[3-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl)oxy]cyclobutoxy]pyridin-2-yl]prop-2-yn-1-yl)oxy]-1,2-oxazol-5-yl]butanoyl)pyrrolidine-2-carboxamide (620 mg) were separated by chiral HPLC under the following conditions: Column: CHIRALPAK IG, 2.0 cm I. D*25 cm (5 um); Mobile phase A: MeOH (8 mmol/L NH3.MeOH), Mobile phase B: DCM, Flow rate: 20 mL/min, Gradient: 30% B to 30% B in 17 min; 220/254 nm; RT1: 8.495 min; RT2: 12.854 min. This yielded 137 mg (22%) of isomer 1, tentatively identified as (2S,4R)-4-hydroxy-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]-1-[(2S)-3-methyl-2-[3-[(3-[5-[(1r,3r)-3-[(4-[3-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl)oxy]cyclobutoxy]pyridin-2-yl]prop-2-yn-1-yl)oxy]-1,2-oxazol-5-yl]butanoyl]pyrrolidine-2-carboxamide (pale yellow solid), and (2S,4R)-4-hydroxy-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]-1-[(2R)-3-methyl-2-[3-[(3-[5-[(1r,3r)-3-[(4-[3-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl]pyridin-2-yl)oxy]cyclobutoxy]pyridin-2-yl]prop-2-yn-1-yl)oxy]-1,2-oxazol-5-yl]butanoyl]pyrrolidine-2-carboxamide (off-white solid) was obtained in an amount of 199 mg (32%).
類似の手順を使用した例示的な化合物226、227、231、232、236、および237。 Exemplary compounds 226, 227, 231, 232, 236, and 237 were synthesized using similar procedures.
例示的な化合物76の例示的な合成
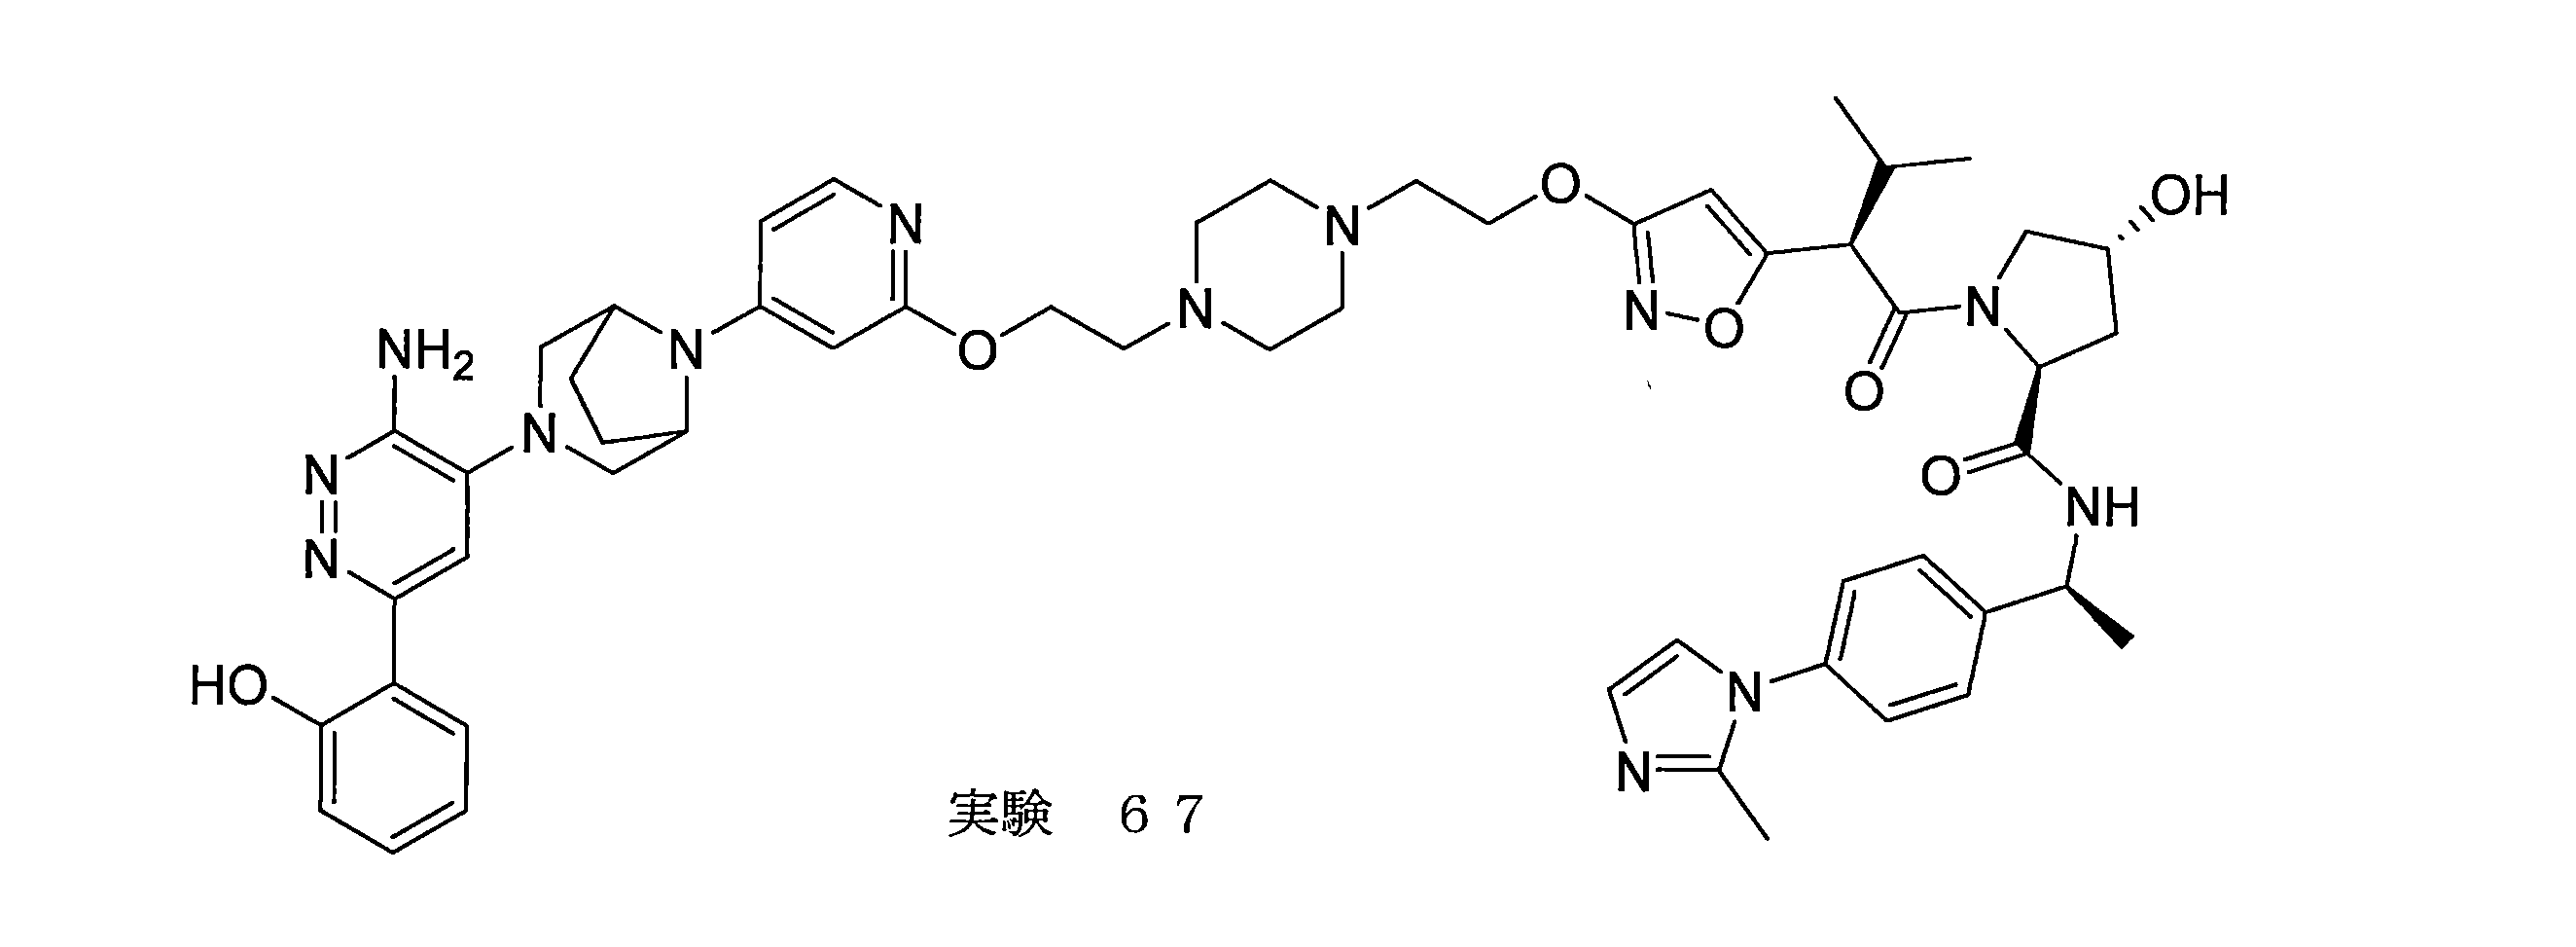
工程1

ジクロロメタン(100mL)中の(1S)-1-(4-ブロモフェニル)エタンアミン(5g、24.99mmol、1.0当量)の溶液に、トリエチルアミン(2.53g、24.99mmol、1.0当量)を0℃で添加した。次いで、混合物を0℃で30分間撹拌した。次いで、ジ-tert-ブチルジカーボネートエステル(5.45g、24.99mmol、1.0当量)を添加し、混合物を、0℃で30分間、続いて20℃で2時間撹拌した。混合物を飽和重炭酸ナトリウム水溶液(20mL)でクエンチし、水(50mL)で希釈し、ジクロロメタン(100mL×2)で抽出し、ブライン(100mL)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、濃縮した。残渣を、石油エーテル:酢酸エチル(50mL、v/v=5/1)で撹拌し、次いで、濾過した。Tert-ブチルN-[(1S)-1-(4-ブロモフェニル)エチル]カルバメート(6.4g、20.68mmol、収率82%)を白色の固体として得た。 To a solution of (1S)-1-(4-bromophenyl)ethanamine (5 g, 24.99 mmol, 1.0 equiv.) in dichloromethane (100 mL) was added triethylamine (2.53 g, 24.99 mmol, 1.0 equiv.) at 0°C. The mixture was then stirred at 0°C for 30 minutes. Di-tert-butyl dicarbonate ester (5.45 g, 24.99 mmol, 1.0 equiv.) was then added and the mixture was stirred at 0°C for 30 minutes followed by 20°C for 2 hours. The mixture was quenched with saturated aqueous sodium bicarbonate (20 mL), diluted with water (50 mL), extracted with dichloromethane (100 mL x 2), washed with brine (100 mL), dried over anhydrous sodium sulfate, filtered and concentrated. The residue was stirred with petroleum ether:ethyl acetate (50 mL, v/v = 5/1) and then filtered. Tert-butyl N-[(1S)-1-(4-bromophenyl)ethyl]carbamate (6.4 g, 20.68 mmol, yield 82%) was obtained as a white solid.
工程2

ジメチルスルホキシド(15mL)中のtert-ブチルN-[(1S)-1-(4-ブロモフェニル)エチル]カルバメート(3g、9.99mmol、1.0当量)および2-メチル-1H-イミダゾール(902mg、10.99mmol、1.1当量)の溶液に、L-プロリン(1.38g、11.99mmol、1.2当量)、炭酸セシウム(9.77g、29.98mmol、3.0当量)、およびヨウ化銅(2.28g、11.99mmol、1.2当量)を窒素下で添加した。次いで、混合物を130℃で12時間撹拌した。混合物を水(200mL)で希釈し、濾過し、次いで、酢酸エチル(200mL×3)で抽出し、ブライン(200mL)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、次いで、濃縮した。残渣を、逆相フラッシュクロマトグラフィー(ギ酸)によって精製した。tert-ブチルN-[(1S)-1-[4-(2-メチルイミダゾール-1-イル)フェニル]エチル]カルバメート(250mg、0.83mmol、収率8%)を黄色の油として得た。 To a solution of tert-butyl N-[(1S)-1-(4-bromophenyl)ethyl]carbamate (3 g, 9.99 mmol, 1.0 eq.) and 2-methyl-1H-imidazole (902 mg, 10.99 mmol, 1.1 eq.) in dimethyl sulfoxide (15 mL), L-proline (1.38 g, 11.99 mmol, 1.2 eq.), cesium carbonate (9.77 g, 29.98 mmol, 3.0 eq.), and copper iodide (2.28 g, 11.99 mmol, 1.2 eq.) were added under nitrogen. The mixture was then stirred at 130° C. for 12 hours. The mixture was diluted with water (200 mL), filtered, and then extracted with ethyl acetate (200 mL×3), washed with brine (200 mL), dried over anhydrous sodium sulfate, filtered, and then concentrated. The residue was purified by reverse-phase flash chromatography (formic acid). tert-Butyl N-[(1S)-1-[4-(2-methylimidazol-1-yl)phenyl]ethyl]carbamate (250 mg, 0.83 mmol, 8% yield) was obtained as a yellow oil.
例示的な化合物47および64について上に記載される手順を使用して、以下のスキームに詳述されるように、tert-ブチルN-[(1S)-1-[4-(2-メチルイミダゾール-1-イル)フェニル]エチル]カルバメートを最終化合物に変換した。

例示的な化合物75を、類似の手順を使用して調製した。 Exemplary compound 75 was prepared using a similar procedure.
例示的な化合物116の例示的な合成
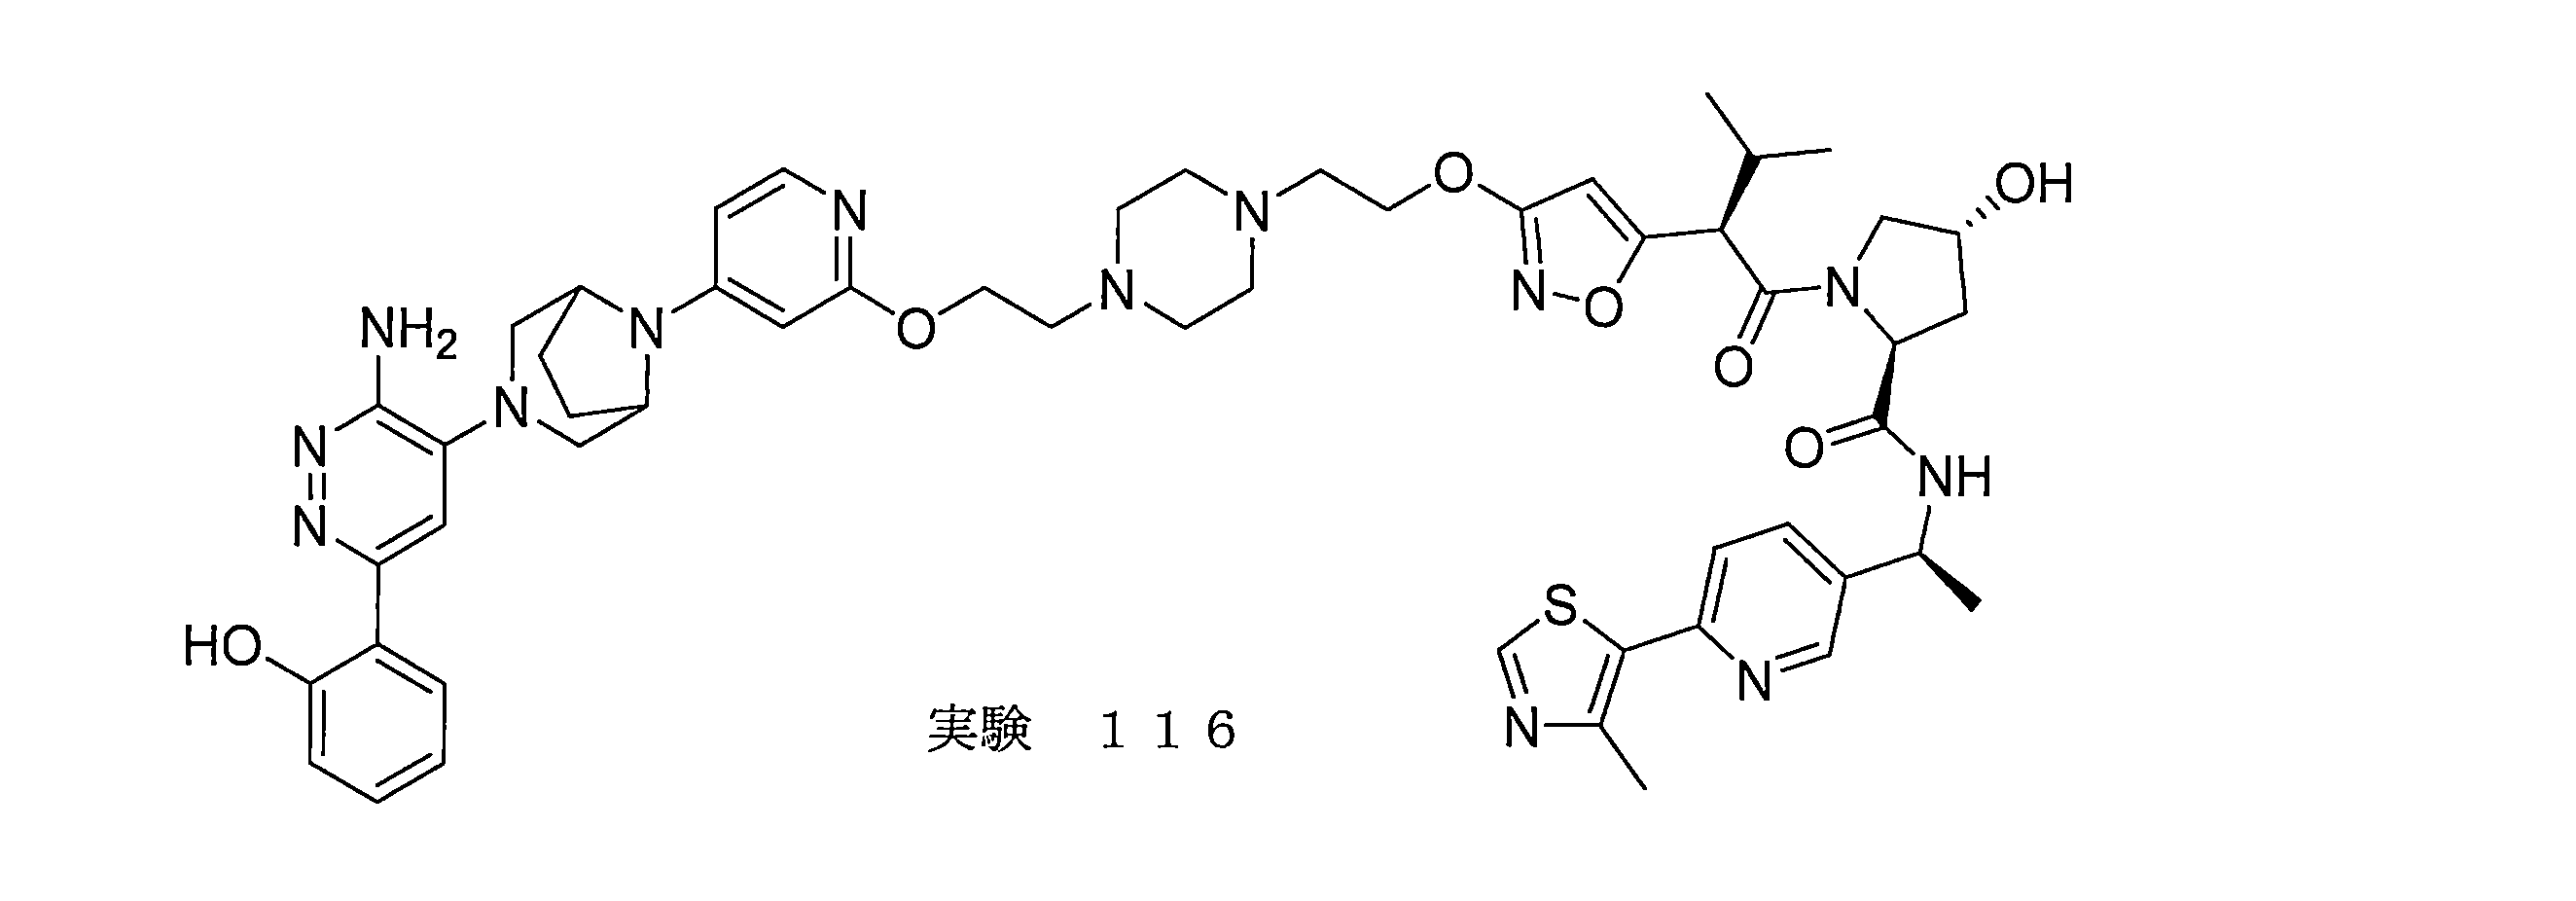
工程1
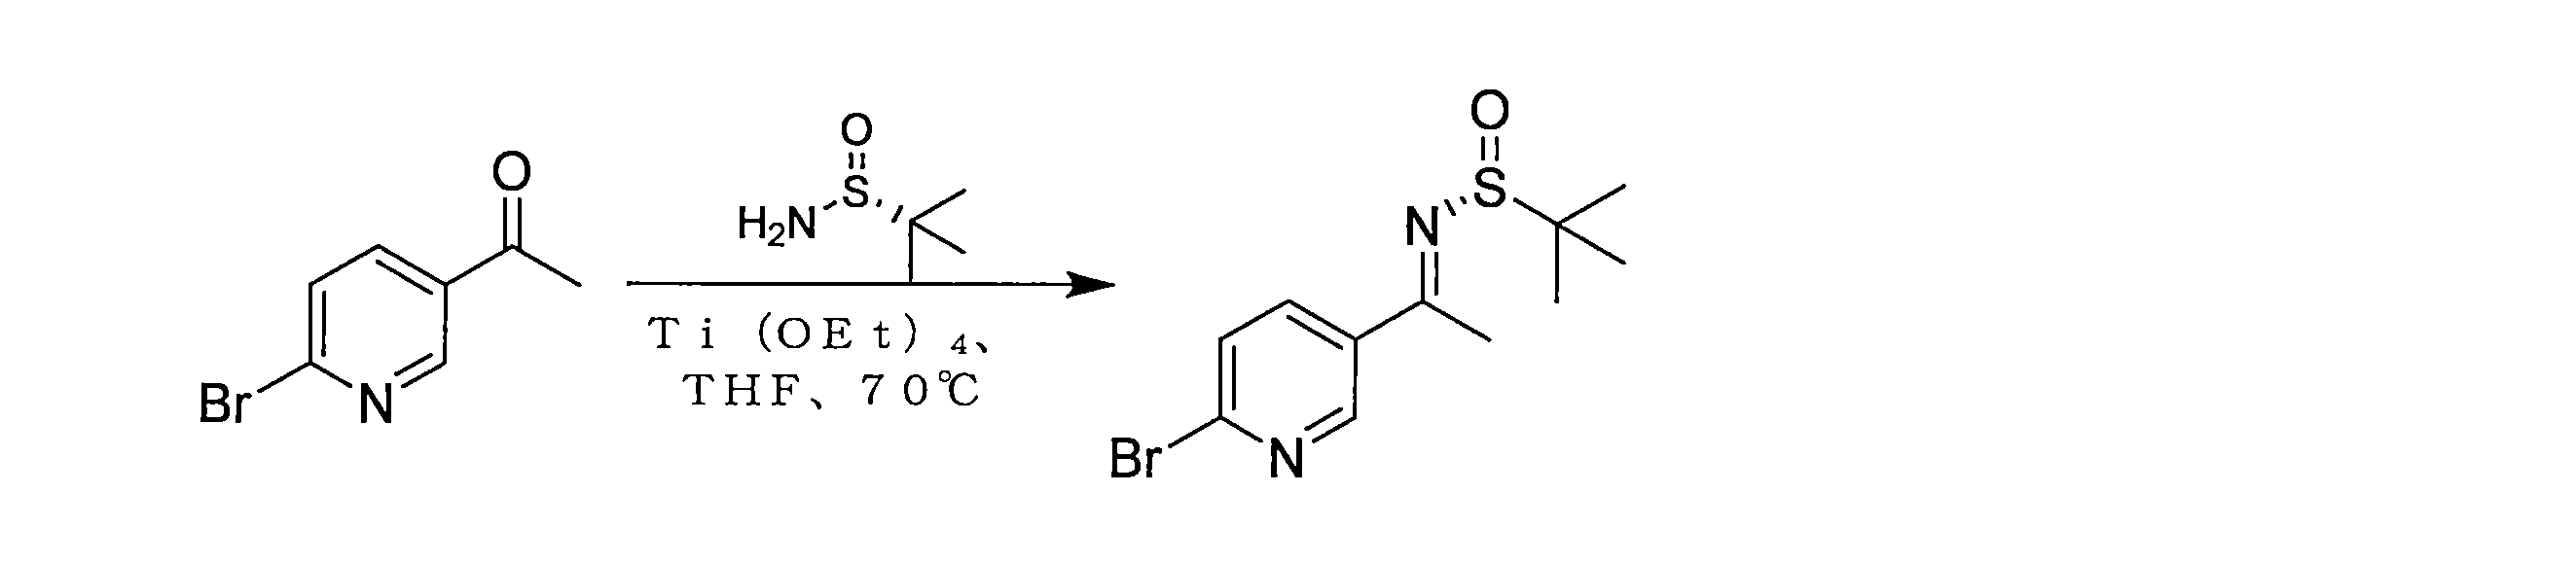
テトラヒドロフラン(50mL)中の1-(6-ブロモ-3-ピリジル)エタノン(11.88g、59.41mmol、1.2当量)およびテトラエトキシチタン(22.58g、99.01mmol、2当量)の溶液に、(R)-2-メチルプロパン-2-スルフィンアミド(6g、49.50mmol、1当量)を窒素下で添加した。反応混合物を70℃で12時間撹拌した。反応混合物を、水(30mL)を添加することによってクエンチし、次いで、水(100mL)でさらに希釈し、濾過し、酢酸エチル(80mL×3)で抽出した。合わせた有機層をブライン(100mL)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、減圧下で濃縮して、所望の化合物(R,E)-N-(1-(6-ブロモピリジン-3-イル)エチリデン)-2-メチルプロパン-2-スルフィンアミド(16g、43.27mmol、収率87%)を淡い黄色の固体として生成した。 To a solution of 1-(6-bromo-3-pyridyl)ethanone (11.88 g, 59.41 mmol, 1.2 eq.) and tetraethoxytitanium (22.58 g, 99.01 mmol, 2 eq.) in tetrahydrofuran (50 mL) was added (R)-2-methylpropane-2-sulfinamide (6 g, 49.50 mmol, 1 eq.) under nitrogen. The reaction mixture was stirred at 70 °C for 12 h. The reaction mixture was quenched by adding water (30 mL) and then further diluted with water (100 mL), filtered, and extracted with ethyl acetate (80 mL x 3). The combined organic layers were washed with brine (100 mL), dried over anhydrous sodium sulfate, filtered, and concentrated under reduced pressure to produce the desired compound (R,E)-N-(1-(6-bromopyridin-3-yl)ethylidene)-2-methylpropane-2-sulfinamide (16 g, 43.27 mmol, 87% yield) as a pale yellow solid.
工程2
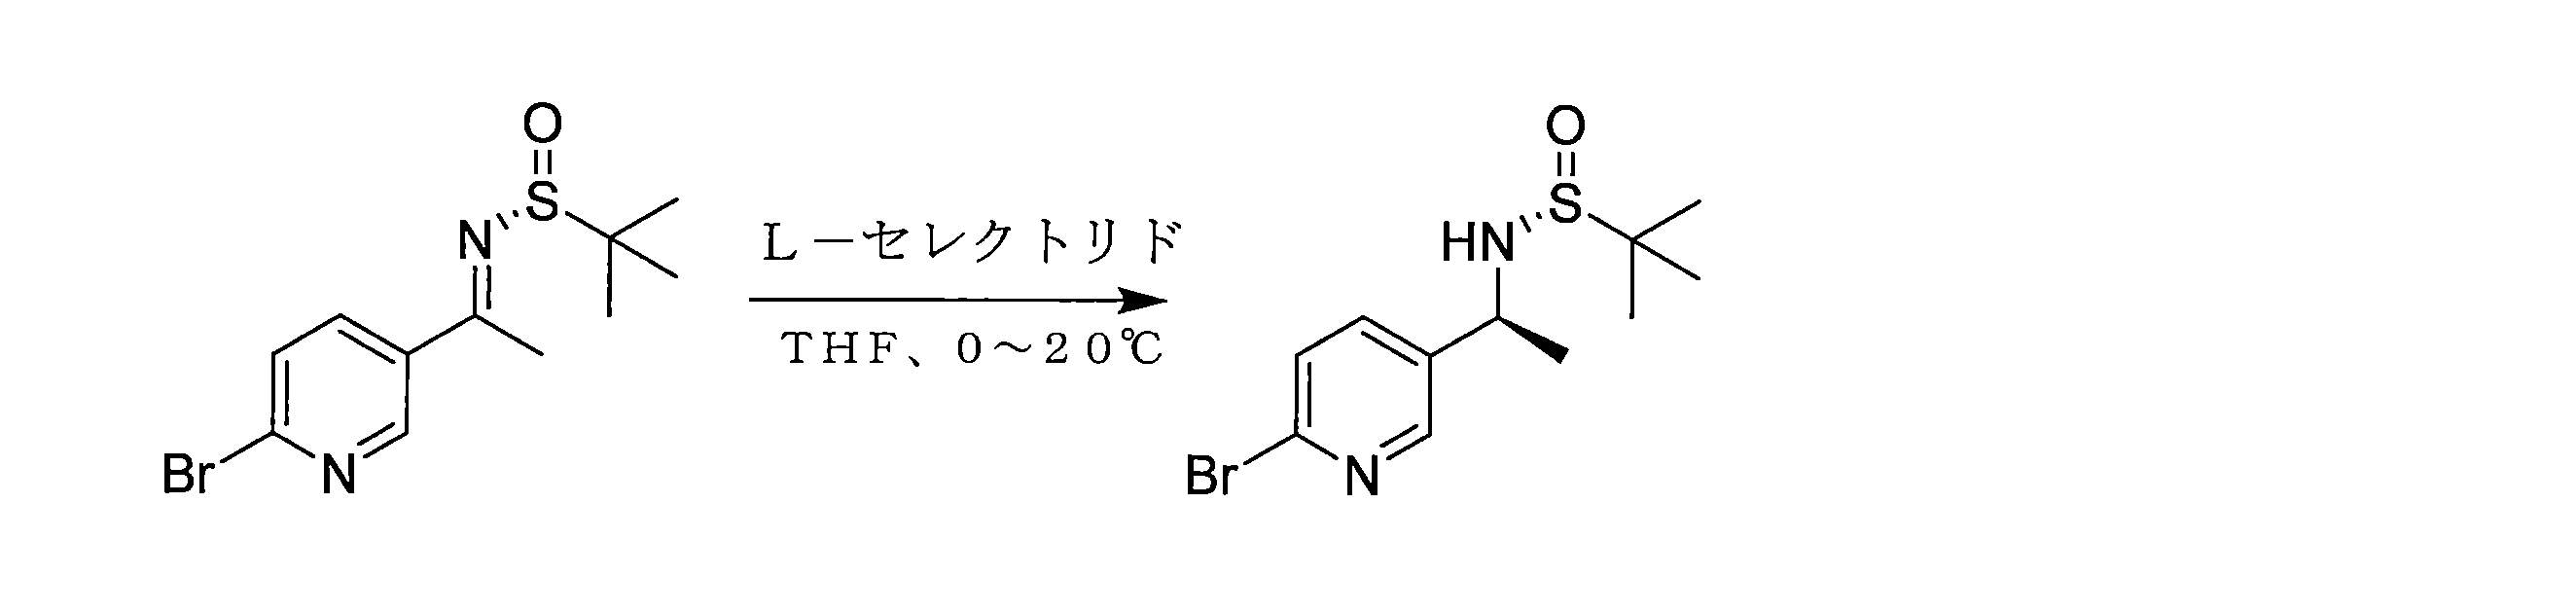
テトラヒドロフラン(160mL)中の(R,E)-N-(1-(6-ブロモピリジン-3-イル)エチリデン)-2-メチルプロパン-2-スルフィンアミド(16g、52.77mmol、1当量)の溶液に、L-セレクトリド(1M、158mL、3当量)を0℃で添加した。反応混合物を温め、20℃で3時間撹拌した。反応混合物を、水(100mL)を添加することによってクエンチし、次いで、水(100mL)でさらに希釈し、濾過し、酢酸エチル(100mL×3)で抽出した。合わせた有機層をブライン(100mL)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、減圧下で濃縮して、粗生成物を得、これを石油エーテル:酢酸エチル=1:1(10mL)からの再結晶化によって精製した。次いで、残渣を、半分取逆相HPLC(FA条件:カラム:Phenomenex Synergi Max-RP 250*50mm*10um;移動相:[水(0.225%FA)-ACN];B%:20%~55%、30分)によって精製した。N-[(1S)-1-(6-ブロモ-3-ピリジル)エチル]-2-メチル-プロパン-2-スルフィンアミド(4.7g、12.89mmol、収率24%)を白色の固体として得た。 To a solution of (R,E)-N-(1-(6-bromopyridin-3-yl)ethylidene)-2-methylpropane-2-sulfinamide (16 g, 52.77 mmol, 1 eq.) in tetrahydrofuran (160 mL) was added L-selectride (1 M, 158 mL, 3 eq.) at 0°C. The reaction mixture was warmed and stirred at 20°C for 3 h. The reaction mixture was quenched by adding water (100 mL), then further diluted with water (100 mL), filtered, and extracted with ethyl acetate (100 mL x 3). The combined organic layers were washed with brine (100 mL), dried over anhydrous sodium sulfate, filtered, and concentrated under reduced pressure to give the crude product, which was purified by recrystallization from petroleum ether:ethyl acetate = 1:1 (10 mL). The residue was then purified by semi-preparative reverse phase HPLC (FA conditions: column: Phenomenex Synergi Max-RP 250*50mm*10um; mobile phase: [water (0.225% FA)-ACN]; B%: 20%-55%, 30 min). N-[(1S)-1-(6-bromo-3-pyridyl)ethyl]-2-methyl-propane-2-sulfinamide (4.7 g, 12.89 mmol, 24% yield) was obtained as a white solid.
工程3
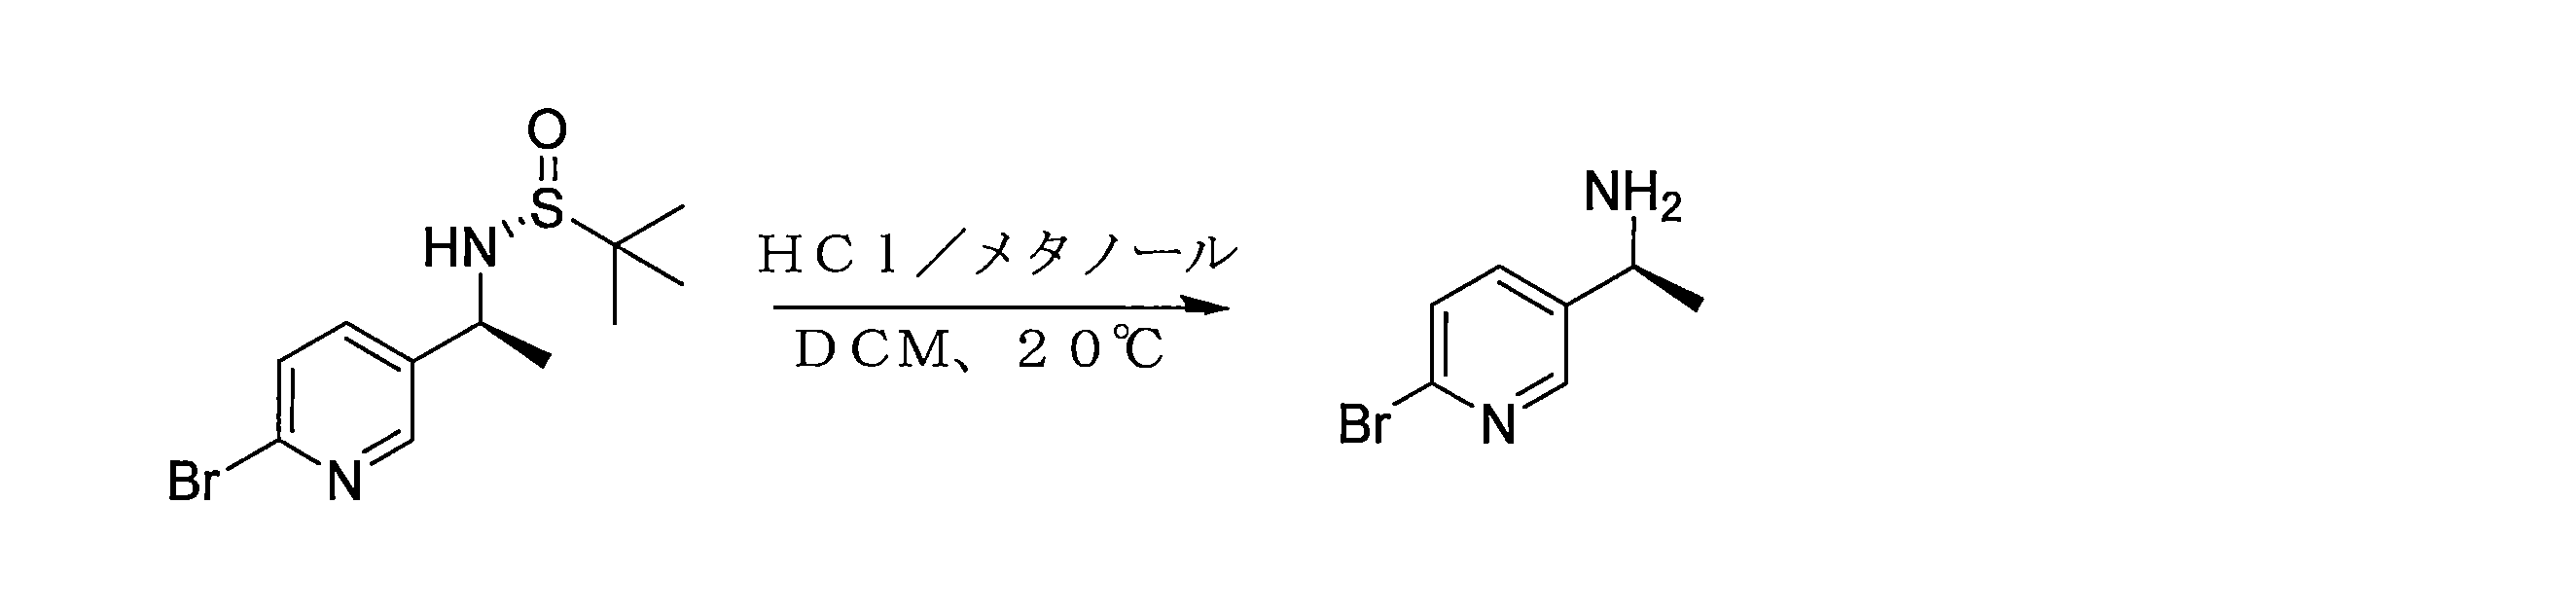
ジクロロメタン(20mL)中のN-[(1S)-1-(6-ブロモ-3-ピリジル)エチル]-2-メチル-プロパン-2-スルフィンアミド(2.6g、8.52mmol、1当量)の溶液に、塩酸/メタノール(4M、21.3mL、10当量)を添加した。反応混合物を20℃で0.15時間撹拌した。反応混合物を減圧下で濃縮して、(1S)-1-(6-ブロモ-3-ピリジル)エタンアミン(1.9g、粗製、塩酸塩)を白色の固体として得た。 To a solution of N-[(1S)-1-(6-bromo-3-pyridyl)ethyl]-2-methyl-propane-2-sulfinamide (2.6 g, 8.52 mmol, 1 equiv.) in dichloromethane (20 mL) was added hydrochloric acid/methanol (4 M, 21.3 mL, 10 equiv.). The reaction mixture was stirred at 20°C for 0.15 h. The reaction mixture was concentrated under reduced pressure to give (1S)-1-(6-bromo-3-pyridyl)ethanamine (1.9 g, crude, hydrochloride salt) as a white solid.
工程4

ジクロロメタン(20mL)中の(1S)-1-(6-ブロモ-3-ピリジル)エタンアミン塩酸塩(1.9g、8.00mmol、1当量)の溶液に、トリエチルアミン(2.43g、24.00mmol、3.34mL、3当量)を0℃で0.5時間、続いてジ-tert-ブチルジカーボネート(2.62g、12.00mmol、2.76mL、1.5当量)を0℃で0.5時間添加した。反応混合物を20℃で12時間撹拌した。反応混合物を、水(20mL)を添加することによってクエンチし、次いで、水60mLでさらに希釈し、酢酸エチル(80mL×3)で抽出した。合わせた有機層をブライン(100mL)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、減圧下で濃縮して、残渣を得、これをシリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=50:1~3:1)によって精製した。tert-ブチルN-[(1S)-1-(6-ブロモ-3-ピリジル)エチル]カルバメート(2.2g、7.06mmol、88%収率)を、白色の固体として得た。 To a solution of (1S)-1-(6-bromo-3-pyridyl)ethanamine hydrochloride (1.9 g, 8.00 mmol, 1 eq.) in dichloromethane (20 mL), triethylamine (2.43 g, 24.00 mmol, 3.34 mL, 3 eq.) was added at 0 °C for 0.5 h, followed by di-tert-butyl dicarbonate (2.62 g, 12.00 mmol, 2.76 mL, 1.5 eq.) at 0 °C for 0.5 h. The reaction mixture was stirred at 20 °C for 12 h. The reaction mixture was quenched by adding water (20 mL), then further diluted with 60 mL of water and extracted with ethyl acetate (80 mL x 3). The combined organic layers were washed with brine (100 mL), dried over anhydrous sodium sulfate, filtered, and concentrated under reduced pressure to give a residue, which was purified by silica gel chromatography (petroleum ether:ethyl acetate = 50:1 to 3:1). tert-Butyl N-[(1S)-1-(6-bromo-3-pyridyl)ethyl]carbamate (2.2 g, 7.06 mmol, 88% yield) was obtained as a white solid.
工程5

ジオキサン(10mL)中の5-ブロモ-4-メチル-チアゾール(800mg、4.49mmol、1当量)および4,4,5,5-テトラメチル-2-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)-1,3,2-ジオキサボロラン(1.37g、5.39mmol、1.2当量)の溶液に、1,1’-ビス(ジフェニルホスフィノ)フェロセン-パラジウム(II)二塩化物(164mg、0.22mmol、0.05当量)および酢酸カリウム(882mg、8.99mmol、2当量)を添加した。反応混合物を100℃で12時間撹拌した。反応混合物を濾過し、減圧下で濃縮して、4-メチル-5-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)チアゾール(1g、粗製)を茶色の油として得、これを精製することなく使用した。 To a solution of 5-bromo-4-methyl-thiazole (800 mg, 4.49 mmol, 1 eq.) and 4,4,5,5-tetramethyl-2-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)-1,3,2-dioxaborolane (1.37 g, 5.39 mmol, 1.2 eq.) in dioxane (10 mL) was added 1,1'-bis(diphenylphosphino)ferrocene-palladium(II) dichloride (164 mg, 0.22 mmol, 0.05 eq.) and potassium acetate (882 mg, 8.99 mmol, 2 eq.). The reaction mixture was stirred at 100 °C for 12 h. The reaction mixture was filtered and concentrated under reduced pressure to give 4-methyl-5-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)thiazole (1 g, crude) as a brown oil, which was used without purification.
工程6

ジオキサン(12mL)および水(2mL)中の4-メチル-5-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)チアゾール(617mg、2.74mmol、1.5当量)およびtert-ブチルN-[(1S)-1-(6-ブロモ-3-ピリジル)エチル]カルバメート(550mg、1.83mmol、1当量)の溶液に、1,1’-ビス(ジフェニルホスフィノ)フェロセン-パラジウム(II)二塩化物(134mg、0.18mmol、0.1当量)および炭酸カリウム(505mg、3.65mmol、2当量)を添加した。反応混合物を100℃で12時間撹拌した。反応混合物を、水(10mL)を添加することによってクエンチし、次いで、水(30mL)で希釈し、濾過し、酢酸エチル(30mL×3)で抽出した。合わせた有機層をブライン(60mL)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、減圧下で濃縮して、残渣を得、これを分取TLC(石油エーテル:酢酸エチル=1:1)によって精製した。所望の化合物tert-ブチルN-[(1S)-1-[6-(4-メチルチアゾール-5-イル)-3-ピリジル]エチル]カルバメート(200mg、0.62mmol、収率34%)を淡い黄色の固体として得た。 To a solution of 4-methyl-5-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)thiazole (617 mg, 2.74 mmol, 1.5 eq.) and tert-butyl N-[(1S)-1-(6-bromo-3-pyridyl)ethyl]carbamate (550 mg, 1.83 mmol, 1 eq.) in dioxane (12 mL) and water (2 mL), 1,1'-bis(diphenylphosphino)ferrocene-palladium(II) dichloride (134 mg, 0.18 mmol, 0.1 eq.) and potassium carbonate (505 mg, 3.65 mmol, 2 eq.) were added. The reaction mixture was stirred at 100 °C for 12 h. The reaction mixture was quenched by adding water (10 mL), then diluted with water (30 mL), filtered and extracted with ethyl acetate (30 mL x 3). The combined organic layers were washed with brine (60 mL), dried over anhydrous sodium sulfate, filtered, and concentrated under reduced pressure to give a residue that was purified by preparative TLC (petroleum ether:ethyl acetate=1:1). The desired compound tert-butyl N-[(1S)-1-[6-(4-methylthiazol-5-yl)-3-pyridyl]ethyl]carbamate (200 mg, 0.62 mmol, 34% yield) was obtained as a pale yellow solid.
例示的な化合物76について上に記載されるように、tert-ブチルN-[(1S)-1-[6-(4-メチルチアゾール-5-イル)-3-ピリジル]エチル]カルバメートを最終化合物に変換した。 tert-Butyl N-[(1S)-1-[6-(4-methylthiazol-5-yl)-3-pyridyl]ethyl]carbamate was converted to the final compound as described above for exemplary compound 76.
例示的な化合物160を、類似の手順を使用して調製した。 Exemplary compound 160 was prepared using a similar procedure.
例示的な化合物117の例示的な合成

工程1
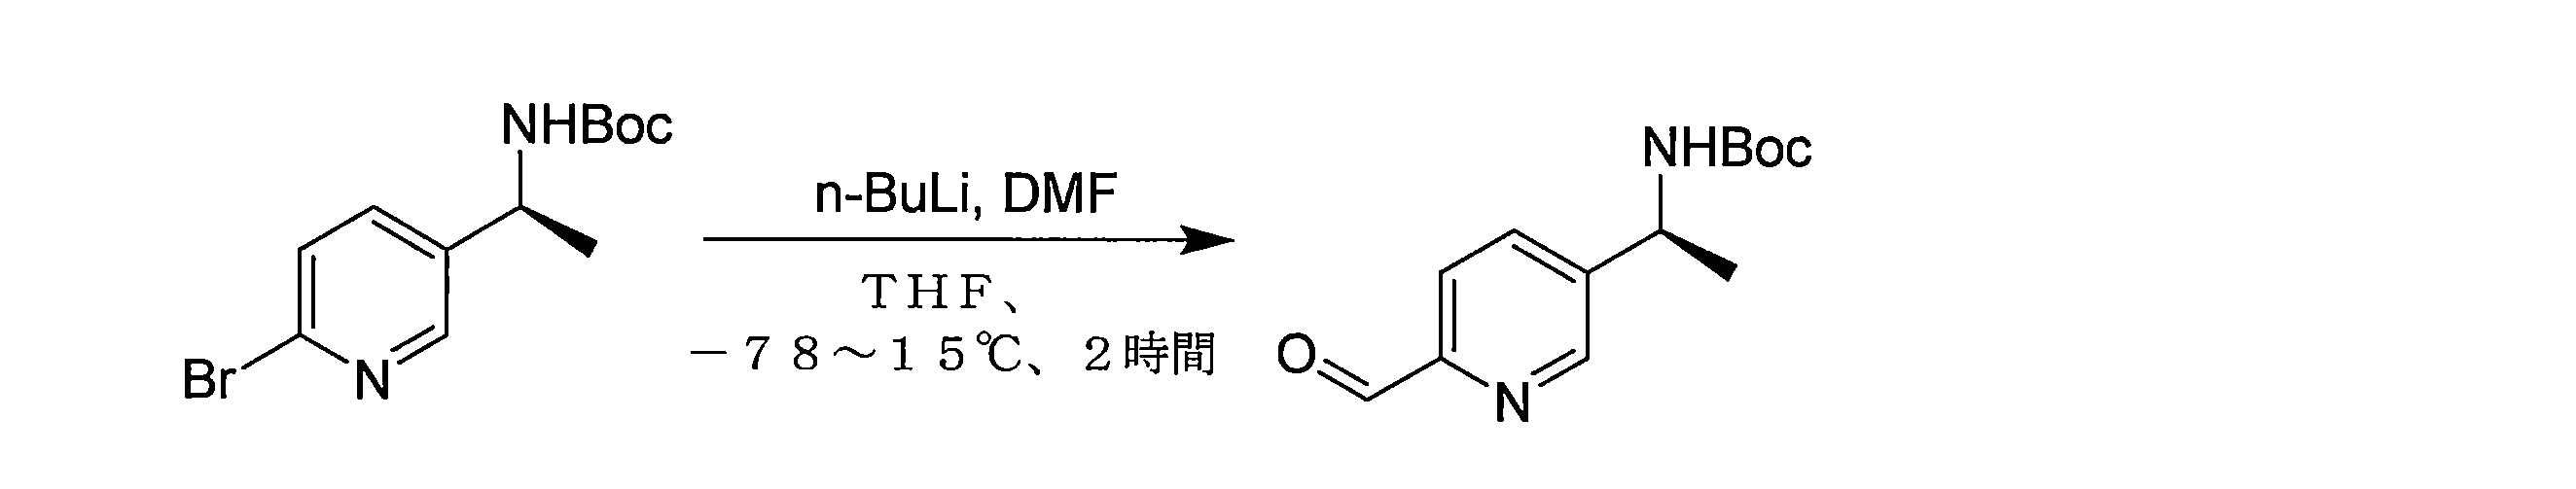
テトラヒドロフラン(10mL)中のtert-ブチルN-[(1S)-1-(6-ブロモ-3-ピリジル)エチル]カルバメート(1g、3.32mmol、1当量)の溶液に、n-ブチルリチウム(2.5M、2.9mL、2.2当量)を-78℃で添加した。混合物を-78℃で0.5時間撹拌し、次いで混合物に、N,N-ジメチルホルムアミド(728mg、9.96mmol、0.8mL、3当量)を-78℃で添加し、混合物を-78℃で0.5時間撹拌した。混合物を15℃まで1時間温めた。反応混合物を、飽和塩化アンモニウム(10mL)を添加することによってクエンチし、次いで、水(20mL)で希釈し、濾過し、酢酸エチル(30mL×3)で抽出した。合わせた有機層をブライン(60mL)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、減圧下で濃縮して、残渣を得た。残渣を、シリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=10:1~1:1)によって精製した。所望の化合物tert-ブチルN-[(1S)-1-(6-ホルミル-3-ピリジル)エチル]カルバメート(550mg、2.20mmol、収率66%)を淡い黄色の固体として得た。 To a solution of tert-butyl N-[(1S)-1-(6-bromo-3-pyridyl)ethyl]carbamate (1 g, 3.32 mmol, 1 eq.) in tetrahydrofuran (10 mL) was added n-butyllithium (2.5 M, 2.9 mL, 2.2 eq.) at -78 °C. The mixture was stirred at -78 °C for 0.5 h, then N,N-dimethylformamide (728 mg, 9.96 mmol, 0.8 mL, 3 eq.) was added to the mixture at -78 °C and the mixture was stirred at -78 °C for 0.5 h. The mixture was warmed to 15 °C for 1 h. The reaction mixture was quenched by adding saturated ammonium chloride (10 mL), then diluted with water (20 mL), filtered, and extracted with ethyl acetate (30 mL x 3). The combined organic layers were washed with brine (60 mL), dried over anhydrous sodium sulfate, filtered, and concentrated under reduced pressure to give a residue. The residue was purified by silica gel chromatography (petroleum ether:ethyl acetate=10:1 to 1:1). The desired compound tert-butyl N-[(1S)-1-(6-formyl-3-pyridyl)ethyl]carbamate (550 mg, 2.20 mmol, 66% yield) was obtained as a pale yellow solid.
工程2

メタノール(10mL)中のtert-ブチルN-[(1S)-1-(6-ホルミル-3-ピリジル)エチル]カルバメート(550mg、2.20mmol、1当量)および1-(1-イソシアノエチルスルホニル)-4-メチル-ベンゼン(506mg、2.42mmol、1.1当量)の溶液に、炭酸カリウム(607mg、4.39mmol、2当量)を添加した。反応混合物を70℃で12時間撹拌した。反応混合物を、水(20mL)を添加することによってクエンチし、次いで、水(10mL)で希釈し、濾過し、酢酸エチル(30mL×3)で抽出した。合わせた有機層をブライン(60mL)で洗浄し、硫酸ナトリウムで乾燥させ、濾過し、減圧下で濃縮して、残渣を得た。残渣を、シリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=50:1~1:1)によって精製した。所望の化合物tert-ブチルN-[(1S)-1-[6-(4-メチルオキサゾール-5-イル)-3-ピリジル]エチル]カルバメート(570mg、1.65mmol、収率75%、純度87%)を白色の固体として得た。 To a solution of tert-butyl N-[(1S)-1-(6-formyl-3-pyridyl)ethyl]carbamate (550 mg, 2.20 mmol, 1 eq.) and 1-(1-isocyanoethylsulfonyl)-4-methyl-benzene (506 mg, 2.42 mmol, 1.1 eq.) in methanol (10 mL), potassium carbonate (607 mg, 4.39 mmol, 2 eq.) was added. The reaction mixture was stirred at 70 °C for 12 h. The reaction mixture was quenched by adding water (20 mL), then diluted with water (10 mL), filtered, and extracted with ethyl acetate (30 mL x 3). The combined organic layers were washed with brine (60 mL), dried over sodium sulfate, filtered, and concentrated under reduced pressure to give a residue. The residue was purified by silica gel chromatography (petroleum ether:ethyl acetate = 50:1 to 1:1). The desired compound tert-butyl N-[(1S)-1-[6-(4-methyloxazol-5-yl)-3-pyridyl]ethyl]carbamate (570 mg, 1.65 mmol, 75% yield, 87% purity) was obtained as a white solid.
例示的な化合物76について記載されるように、tert-ブチルN-[(1S)-1-[6-(4-メチルオキサゾール-5-イル)-3-ピリジル]エチル]カルバメートを最終化合物に変換した。 tert-Butyl N-[(1S)-1-[6-(4-methyloxazol-5-yl)-3-pyridyl]ethyl]carbamate was converted to the final compound as described for exemplary compound 76.
例示的な化合物161、229、および230を、類似の手順を使用して調製した。 Exemplary compounds 161, 229, and 230 were prepared using similar procedures.
例示的な化合物81の例示的な合成

工程1
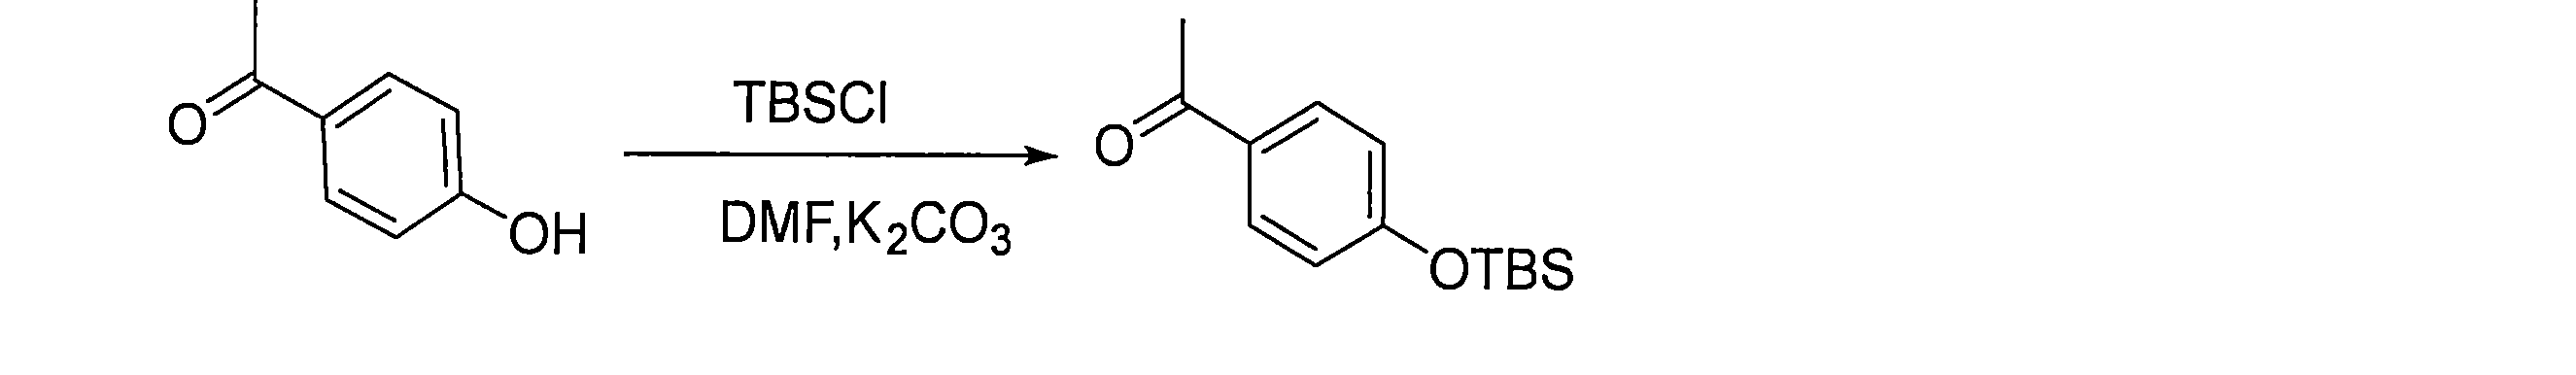
250mLの丸底フラスコに、DMF(100mL)中の1-(4-ヒドロキシフェニル)エタン-1-オン(6.0g、44.08mmol、1当量)、tert-ブチル(クロロ)ジメチルシラン(10g、66.1mmol、1.5当量)、およびK2CO3(12g、88.14mmol、2当量)を入れた。得られた混合物を室温で一晩撹拌した。次いで、反応物を、水(100mL)を添加することによってクエンチした。得られた混合物をジクロロメタン(100mL×3)で抽出し、有機層を合わせ、無水硫酸ナトリウム上で乾燥させ、減圧下で濃縮した。残渣を、酢酸エチル/石油エーテル(1:10)で溶出するシリカゲルカラム上に適用した。これにより、5.2g(47%)の1-[4-[(tert-ブチルジメチルシリル)オキシ]フェニル]エタン-1-オンを灰色がかった白色の固体として得た。 A 250 mL round bottom flask was charged with 1-(4-hydroxyphenyl)ethan-1-one (6.0 g, 44.08 mmol, 1 equiv.), tert-butyl(chloro)dimethylsilane (10 g, 66.1 mmol, 1.5 equiv.), and K2CO3 (12 g, 88.14 mmol, 2 equiv.) in DMF (100 mL). The resulting mixture was stirred at room temperature overnight. The reaction was then quenched by adding water (100 mL). The resulting mixture was extracted with dichloromethane (100 mL x 3), and the organic layers were combined, dried over anhydrous sodium sulfate, and concentrated under reduced pressure. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (1:10). This gave 5.2 g (47%) of 1-[4-[(tert-butyldimethylsilyl)oxy]phenyl]ethan-1-one as an off-white solid.
工程2

250mLの丸底フラスコに、MeOH(100mL)中の1-[4-[(tert-ブチルジメチルシリル)オキシ]フェニル]エタン-1-オン(5g、20.0mmol、1当量)を入れ、それに水素化ホウ素ナトリウム(1.5g、39.94mmol、2当量)を室温で少しずつ添加した。得られた溶液を室温で5時間撹拌し、次いで、減圧下で濃縮した。残渣を、酢酸エチル/石油エーテル(1/1)で溶出するシリカゲルカラム上に適用した。これにより、3.8g(76%)の1-[4-[(tert-ブチルジメチルシリル)オキシ]フェニル]エタン-1-オールを黄色の油として得た。 A 250 mL round bottom flask was charged with 1-[4-[(tert-butyldimethylsilyl)oxy]phenyl]ethan-1-one (5 g, 20.0 mmol, 1 equiv.) in MeOH (100 mL), to which sodium borohydride (1.5 g, 39.94 mmol, 2 equiv.) was added portionwise at room temperature. The resulting solution was stirred at room temperature for 5 h and then concentrated under reduced pressure. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (1/1). This gave 3.8 g (76%) of 1-[4-[(tert-butyldimethylsilyl)oxy]phenyl]ethan-1-ol as a yellow oil.
工程3

250mLの丸底フラスコに、ジクロロメタン(100mL)中の1-[4-[(tert-ブチルジメチルシリル)オキシ]フェニル]エタン-1-オール(3.8g、15.05mmol、1当量)およびTEA(6.1g、60.21mmol、4当量)を入れ、それにメタンスルホニルクロリド(2.1g、18.07mmol、1.20当量)を室温でゆっくり添加した。得られた混合物を、室温で2時間撹拌した。混合物を減圧下で濃縮した。これにより、3.6gの粗1-[4-[(tert-ブチルジメチルシリル)オキシ]フェニル]エチルメタンスルホネートを黄色の固体として得た。 A 250 mL round bottom flask was charged with 1-[4-[(tert-butyldimethylsilyl)oxy]phenyl]ethan-1-ol (3.8 g, 15.05 mmol, 1 eq.) and TEA (6.1 g, 60.21 mmol, 4 eq.) in dichloromethane (100 mL), to which methanesulfonyl chloride (2.1 g, 18.07 mmol, 1.20 eq.) was added slowly at room temperature. The resulting mixture was stirred at room temperature for 2 hours. The mixture was concentrated under reduced pressure. This gave 3.6 g of crude 1-[4-[(tert-butyldimethylsilyl)oxy]phenyl]ethyl methanesulfonate as a yellow solid.
工程4
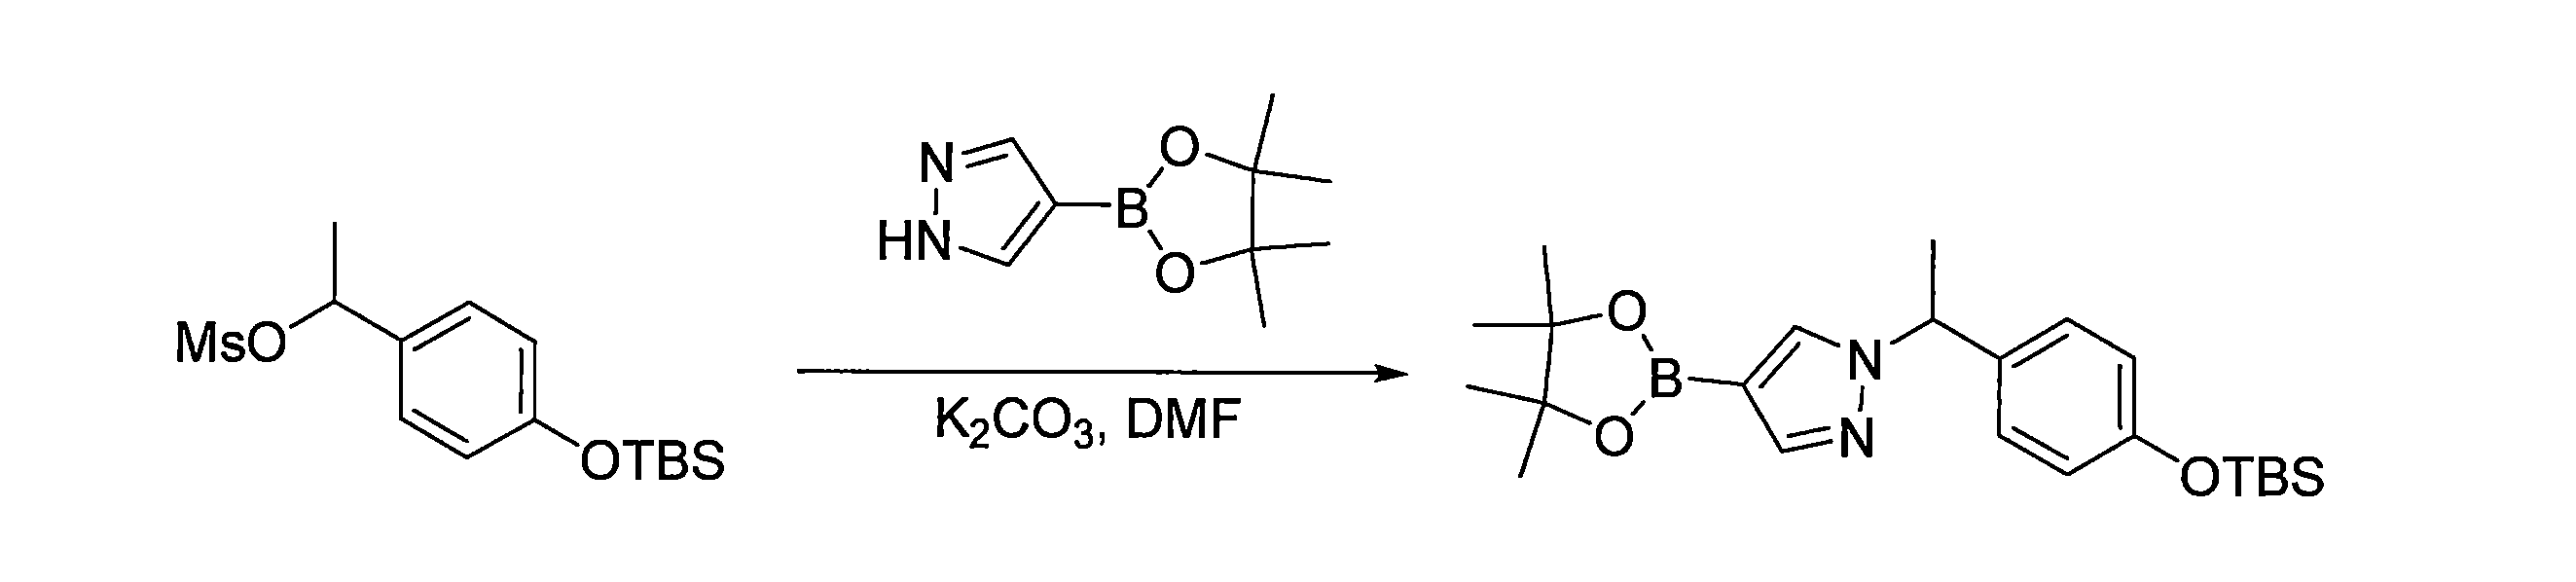
250mLの丸底フラスコに、DMF(150mL)中の1-[4-[(tert-ブチルジメチルシリル)オキシ]フェニル]エチルメタンスルホネート(5.0g、15.13mmol、1当量)、2-(シクロペンタ-1,4-ジエン-1-イル)-4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン(3.5g、18.17mmol、1.20当量)、およびK2CO3(6.3g、45.38mmol、3当量)を窒素雰囲気下で入れた。得られた混合物を90℃で一晩撹拌した。次いで、反応物を、水(150mL)を添加することによってクエンチした。得られた混合物をジクロロメタン(150mL×3)で抽出し、有機層を合わせ、無水硫酸ナトリウムで乾燥させ、減圧下で濃縮した。残渣を、酢酸エチル/石油エーテル(1:5)で溶出するシリカゲルカラム上に適用した。これにより、600mg(9%)の1-(1-[4-[(tert-ブチルジメチルシリル)オキシ]フェニル]エチル)-4-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)-1H-ピラゾールを黄色の固体として得た。 A 250 mL round bottom flask was charged with 1-[4-[(tert-butyldimethylsilyl)oxy]phenyl]ethyl methanesulfonate (5.0 g, 15.13 mmol, 1 eq.), 2-(cyclopenta-1,4-dien-1-yl)-4,4,5,5-tetramethyl-1,3,2-dioxaborolane (3.5 g, 18.17 mmol, 1.20 eq.), and K2CO3 (6.3 g, 45.38 mmol, 3 eq.) in DMF (150 mL) under nitrogen atmosphere. The resulting mixture was stirred at 90 °C overnight. The reaction was then quenched by adding water (150 mL). The resulting mixture was extracted with dichloromethane (150 mL × 3), the organic layers were combined, dried over anhydrous sodium sulfate, and concentrated under reduced pressure. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (1:5). This gave 600 mg (9%) of 1-(1-[4-[(tert-butyldimethylsilyl)oxy]phenyl]ethyl)-4-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)-1H-pyrazole as a yellow solid.
他の実施例について記載されるものと類似の手順を使用して、以下のスキームに従って、1-(1-[4-[(tert-ブチルジメチルシリル)オキシ]フェニル]エチル)-4-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)-1H-ピラゾールを最終化合物に変換した。

例示的な化合物156を、類似の手順を使用して調製した。 Exemplary compound 156 was prepared using a similar procedure.
例示的な化合物162の例示的な合成

工程1

250mLの丸底フラスコに、EtOH(60mL)中の1-(6-フルオロピリジン-3-イル)エタン-1-オン(4.17g、29.97mmol、1当量)の溶液を入れた。溶液を水/氷浴中で0℃に冷却し、H2O(10mL)中の水素化ホウ素ナトリウム(2.27g、61.65mmol、2.06当量)の溶液を滴下で添加した。得られた溶液を室温で0.5時間撹拌した。反応混合物を水(400mL)で慎重にクエンチし、酢酸エチル(200mL×3)で抽出した。有機層を合わせ、無水硫酸ナトリウムで乾燥させ、真空下で濃縮した。残渣を、酢酸エチル/石油エーテル(1:1)で溶出するシリカゲルカラム上に適用した。これにより、3.87g(91%)の1-(6-フルオロピリジン-3-イル)エタン-1-オールを淡い黄色の油として得た。 A 250 mL round bottom flask was charged with a solution of 1-(6-fluoropyridin-3-yl)ethan-1-one (4.17 g, 29.97 mmol, 1 equiv.) in EtOH (60 mL). The solution was cooled to 0° C. in a water/ice bath and a solution of sodium borohydride (2.27 g, 61.65 mmol, 2.06 equiv.) in H2O (10 mL) was added dropwise. The resulting solution was stirred at room temperature for 0.5 h. The reaction mixture was carefully quenched with water (400 mL) and extracted with ethyl acetate (200 mL×3). The organic layers were combined, dried over anhydrous sodium sulfate and concentrated under vacuum. The residue was applied onto a silica gel column eluting with ethyl acetate/petroleum ether (1:1). This gave 3.87 g (91%) of 1-(6-fluoropyridin-3-yl)ethan-1-ol as a pale yellow oil.
工程2

窒素不活性雰囲気下でパージし、維持した500mLの三口丸底フラスコに、ジクロロメタン(200mL)中の1-(6-フルオロピリジン-3-イル)エタン-1-オール(4.06g、28.76mmol、1当量)を入れた。溶液を水/氷浴中で0℃に冷却し、PPh3(11.28g、43.01mmol、1.50当量)およびNBS(7.66g、43.04mmol、1.50当量)を0℃で逐次的に添加した。得られた溶液を室温で2時間撹拌した。反応混合物を真空下で濃縮した。残渣を、酢酸エチル/石油エーテル(1:9)で溶出するシリカゲルカラム上に適用した。これにより、5.83g(99%)の5-(1-ブロモエチル)-2-フルオロピリジンを淡い黄色の液体として得た。 A 500 mL three-neck round bottom flask purged and maintained under nitrogen inert atmosphere was charged with 1-(6-fluoropyridin-3-yl)ethan-1-ol (4.06 g, 28.76 mmol, 1 equiv.) in dichloromethane (200 mL). The solution was cooled to 0 °C in a water/ice bath and PPh3 (11.28 g, 43.01 mmol, 1.50 equiv.) and NBS (7.66 g, 43.04 mmol, 1.50 equiv.) were added sequentially at 0 °C. The resulting solution was stirred at room temperature for 2 h. The reaction mixture was concentrated under vacuum. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (1:9). This gave 5.83 g (99%) of 5-(1-bromoethyl)-2-fluoropyridine as a pale yellow liquid.
工程3
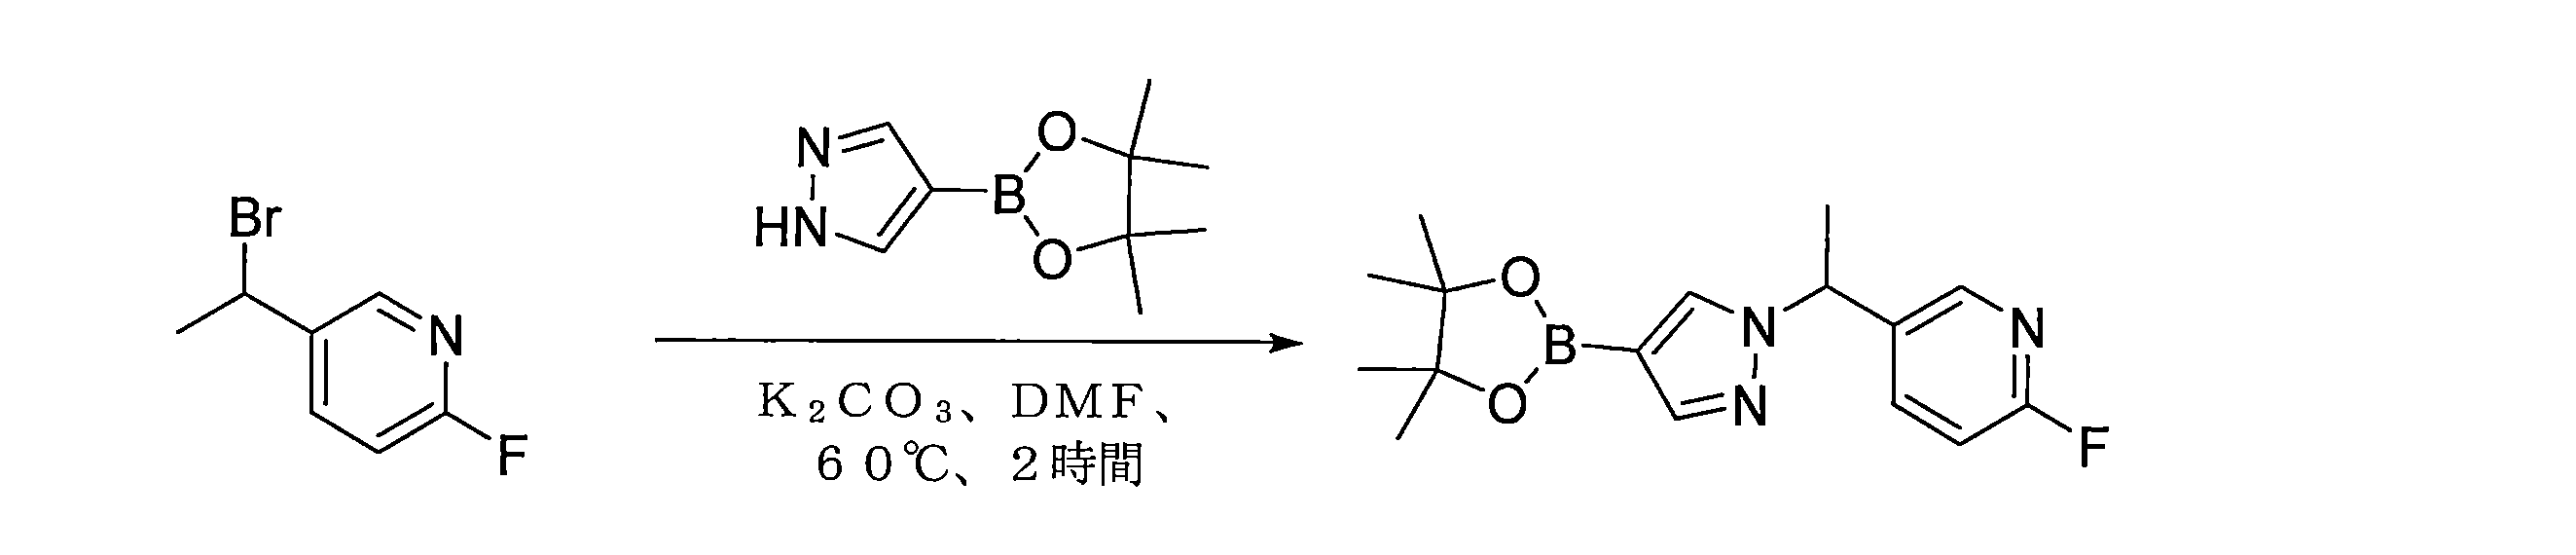
100mLの丸底フラスコに、DMF(50mL)中の5-(1-ブロモエチル)-2-フルオロピリジン(3.06g、15.00mmol、1当量)、4-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)-1H-ピラゾール(3.20g、16.49mmol、1.10当量)、およびK2CO3(4.15g、30.03mmol、2.00当量)の溶液を入れた。得られた混合物を、油浴中で、60℃で2時間撹拌した。反応混合物を室温に冷却し、水(400mL)で希釈した。混合物を酢酸エチル(100mL×4)で抽出し、合わせた有機層をブライン(300mL×2)で洗浄し、無水硫酸ナトリウムで乾燥させ、真空下で濃縮した。これにより、3.43g(72%)の2-フルオロ-5-[1-[4-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)-1H-ピラゾール-1-イル]エチル]ピリジンを淡い黄色の固体として得た。 A 100 mL round-bottom flask was charged with a solution of 5-(1-bromoethyl)-2-fluoropyridine (3.06 g, 15.00 mmol, 1 equiv.), 4-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)-1H-pyrazole (3.20 g, 16.49 mmol, 1.10 equiv.), and K2CO3 (4.15 g, 30.03 mmol, 2.00 equiv.) in DMF (50 mL). The resulting mixture was stirred in an oil bath at 60 °C for 2 h. The reaction mixture was cooled to room temperature and diluted with water (400 mL). The mixture was extracted with ethyl acetate (100 mL × 4) and the combined organic layers were washed with brine (300 mL × 2), dried over anhydrous sodium sulfate, and concentrated under vacuum. This yielded 3.43 g (72%) of 2-fluoro-5-[1-[4-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)-1H-pyrazol-1-yl]ethyl]pyridine as a pale yellow solid.
工程4

窒素不活性雰囲気下でパージし、維持した100mLの丸底フラスコに、ジオキサン(50mL)およびH2O(8mL)中の2-フルオロ-5-[1-[4-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)-1H-ピラゾール-1-イル]エチル]ピリジン(2.54g、8.01mmol、1当量、4-ブロモ-6-クロロピリダジン-3-アミン(2.00g、9.60mmol、1.20当量)、Na2CO3(2.12g、20.00mmol、2.50当量)、およびPd(dppf)Cl2.CH2Cl2(327mg、0.40mmol、0.05当量)を入れた。得られた混合物を、油浴中で90℃において2時間撹拌した。反応混合物を、室温に冷却し、水(200mL)で希釈した。混合物を酢酸エチル(300mL×2)で抽出し、有機層を合わせ、無水硫酸ナトリウム上で乾燥させ、真空下で濃縮した。残渣を、ジクロロメタン/メタノール(20:1)で溶出するシリカゲルカラム上に適用した。これにより、1.88g(74%)の6-クロロ-4-[1-[1-(6-フルオロピリジン-3-イル)エチル]-1H-ピラゾール-4-イル]ピリダジン-3-アミンを茶色の固体として得た。 In a 100 mL round bottom flask purged and maintained under a nitrogen inert atmosphere, 2-fluoro-5-[1-[4-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)-1H-pyrazol-1-yl]ethyl]pyridine (2.54 g, 8.01 mmol, 1 equiv., 4-bromo-6-chloropyridazin-3-amine (2.00 g, 9.60 mmol, 1.20 equiv.), Na2CO3 (2.12 g, 20.00 mmol, 2.50 equiv.), and Pd(dppf)Cl2.CH2Cl2 (327 mg, 0.40 mmol) in dioxane (50 mL) and H2O (8 mL) were added. mol, 0.05 equiv.) was charged. The resulting mixture was stirred in an oil bath at 90° C. for 2 hours. The reaction mixture was cooled to room temperature and diluted with water (200 mL). The mixture was extracted with ethyl acetate (300 mL×2), and the organic layers were combined, dried over anhydrous sodium sulfate, and concentrated under vacuum. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (20:1). This gave 1.88 g (74%) of 6-chloro-4-[1-[1-(6-fluoropyridin-3-yl)ethyl]-1H-pyrazol-4-yl]pyridazin-3-amine as a brown solid.
工程5

窒素不活性雰囲気でパージし、維持した30mLの密封チューブに、ジオキサン(18mL)およびH2O(3mL)中の6-クロロ-4-[1-[1-(6-フルオロピリジン-3-イル)エチル]-1H-ピラゾール-4-イル]ピリダジン-3-アミン(765mg、2.40mmol、1当量)、[2-(メトキシメトキシ)フェニル]ボロン酸(524mg、2.88mmol、1.20当量)、K2CO3(829mg、6.00mmol、2.50当量)、およびPd(dppf)Cl2.CH2Cl2(196mg、0.24mmol、0.10当量)を入れた。得られた混合物を、油浴中で、100℃で2時間撹拌した。反応混合物を室温に冷却し、真空下で濃縮した。残渣を、ジクロロメタン/メタノール(20:1)で溶出するシリカゲルカラム上に適用した。これにより、860mg(85%)の4-[1-[1-(6-フルオロピリジン-3-イル)エチル]-1H-ピラゾール-4-イル]-6-[2-(メトキシメトキシ)フェニル]ピリダジン-3-アミンを茶色の固体として得た。 In a 30 mL sealed tube purged and maintained with nitrogen inert atmosphere, 6-chloro-4-[1-[1-(6-fluoropyridin-3-yl)ethyl]-1H-pyrazol-4-yl]pyridazin-3-amine (765 mg, 2.40 mmol, 1 equiv.), [2-(methoxymethoxy)phenyl]boronic acid (524 mg, 2.88 mmol, 1.20 equiv.), K2CO3 (829 mg, 6.00 mmol, 2.50 equiv.), and Pd(dppf)Cl2. CH2Cl2 (196 mg, 0.24 mmol, 0.10 equiv.) in dioxane (18 mL) and H2O (3 mL) were placed. The resulting mixture was stirred in an oil bath at 100 °C for 2 h. The reaction mixture was cooled to room temperature and concentrated under vacuum. The residue was applied onto a silica gel column eluting with dichloromethane/methanol (20:1). This yielded 860 mg (85%) of 4-[1-[1-(6-fluoropyridin-3-yl)ethyl]-1H-pyrazol-4-yl]-6-[2-(methoxymethoxy)phenyl]pyridazin-3-amine as a brown solid.
工程6
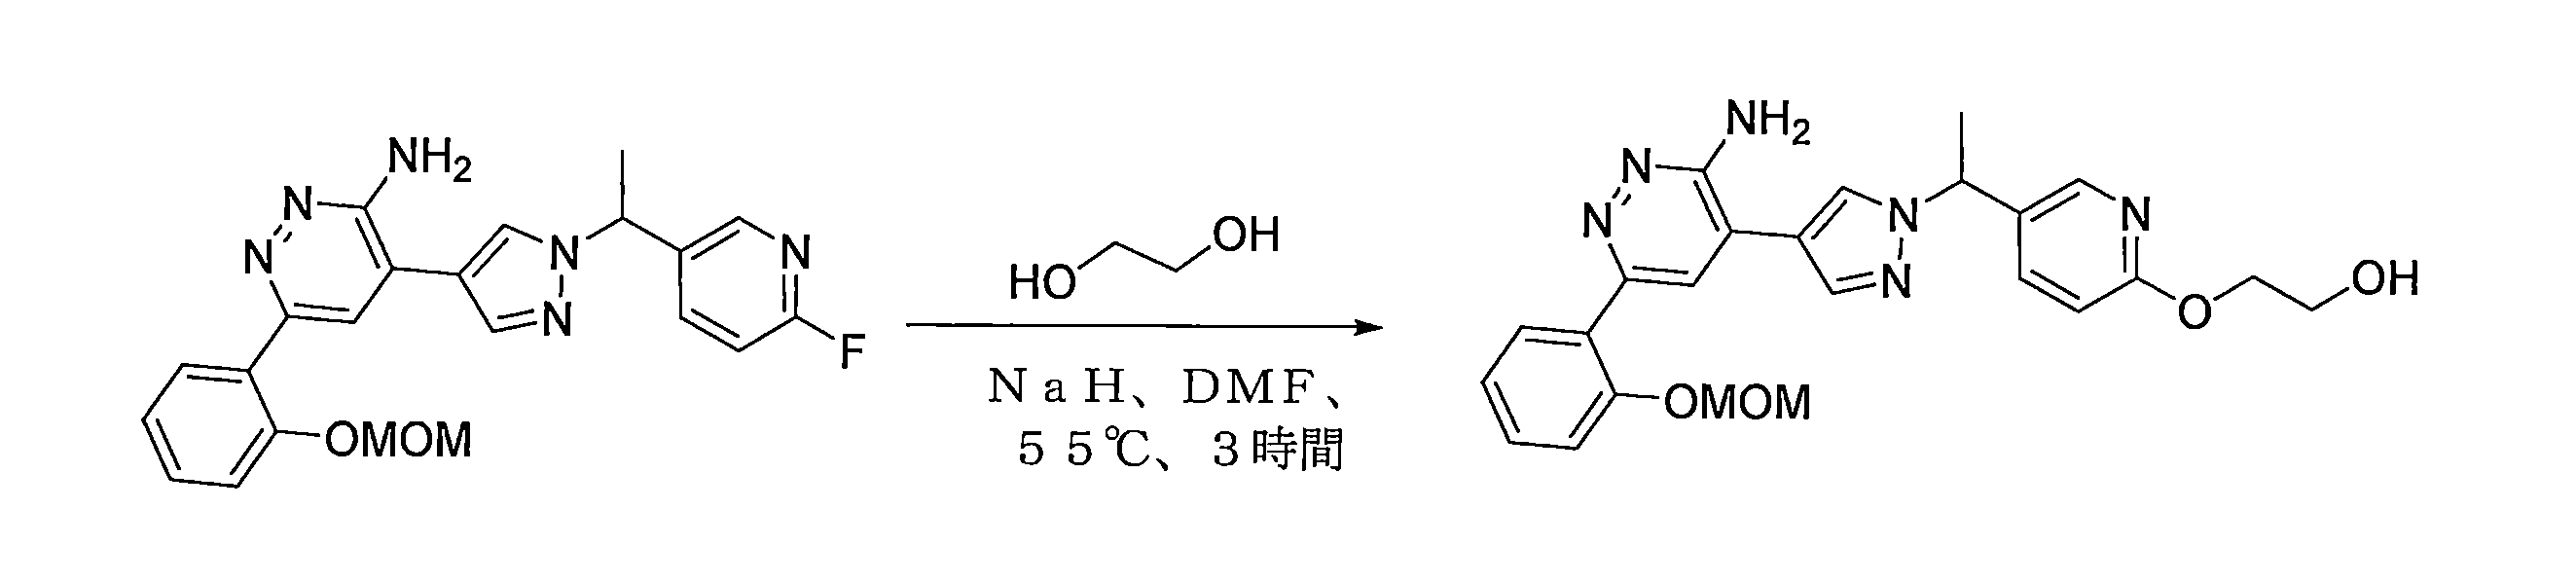
窒素不活性雰囲気下でパージし、維持した50mLの丸底フラスコに、DMF(15mL)中のエタン-1,2-ジオール(1.48g、23.84mmol、10.03当量)を入れた。溶液を水/氷浴中で0℃に冷却し、これにNaH(0.76g、19.00mmol、7.99当量、60%)を0℃で少しずつ添加した。得られた混合物を室温で1.5時間撹拌し、その後、DMF(5mL)中の4-[1-[1-(6-フルオロピリジン-3-イル)エチル]-1H-ピラゾール-4-イル]-6-[2-(メトキシメトキシ)フェニル]ピリダジン-3-アミン(1.00g、2.38mmol、1当量)の溶液を0℃で、滴下で添加した。温度を油浴中で、55℃で維持しながら、反応混合物をさらに3時間撹拌した。反応混合物を室温に冷却し、次いで、水(200mL)を添加することによってクエンチした。得られた混合物を酢酸エチル(300mL×2)で抽出した。合わせた有機層をブライン(200mL×2)で洗浄し、無水硫酸ナトリウム上で乾燥させ、真空下で濃縮させた。残渣を、ジクロロメタン/メタノール(15:1)で溶出するシリカゲルカラム上に適用した。これにより、361mg(33%)の2-([5-[1-(4-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-1H-ピラゾール-1-イル)エチル]ピリジン-2-イル]オキシ)エタン-1-オールを淡い黄色の固体として得た。 A 50 mL round bottom flask purged and maintained under a nitrogen inert atmosphere was charged with ethane-1,2-diol (1.48 g, 23.84 mmol, 10.03 equiv.) in DMF (15 mL). The solution was cooled to 0°C in a water/ice bath, to which NaH (0.76 g, 19.00 mmol, 7.99 equiv., 60%) was added portionwise at 0°C. The resulting mixture was stirred at room temperature for 1.5 h, after which a solution of 4-[1-[1-(6-fluoropyridin-3-yl)ethyl]-1H-pyrazol-4-yl]-6-[2-(methoxymethoxy)phenyl]pyridazin-3-amine (1.00 g, 2.38 mmol, 1 equiv.) in DMF (5 mL) was added dropwise at 0°C. The reaction mixture was stirred for an additional 3 h while the temperature was maintained at 55°C in an oil bath. The reaction mixture was cooled to room temperature and then quenched by adding water (200 mL). The resulting mixture was extracted with ethyl acetate (300 mL x 2). The combined organic layers were washed with brine (200 mL x 2), dried over anhydrous sodium sulfate, and concentrated under vacuum. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (15:1). This gave 361 mg (33%) of 2-([5-[1-(4-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-1H-pyrazol-1-yl)ethyl]pyridin-2-yl]oxy)ethan-1-ol as a pale yellow solid.
工程7
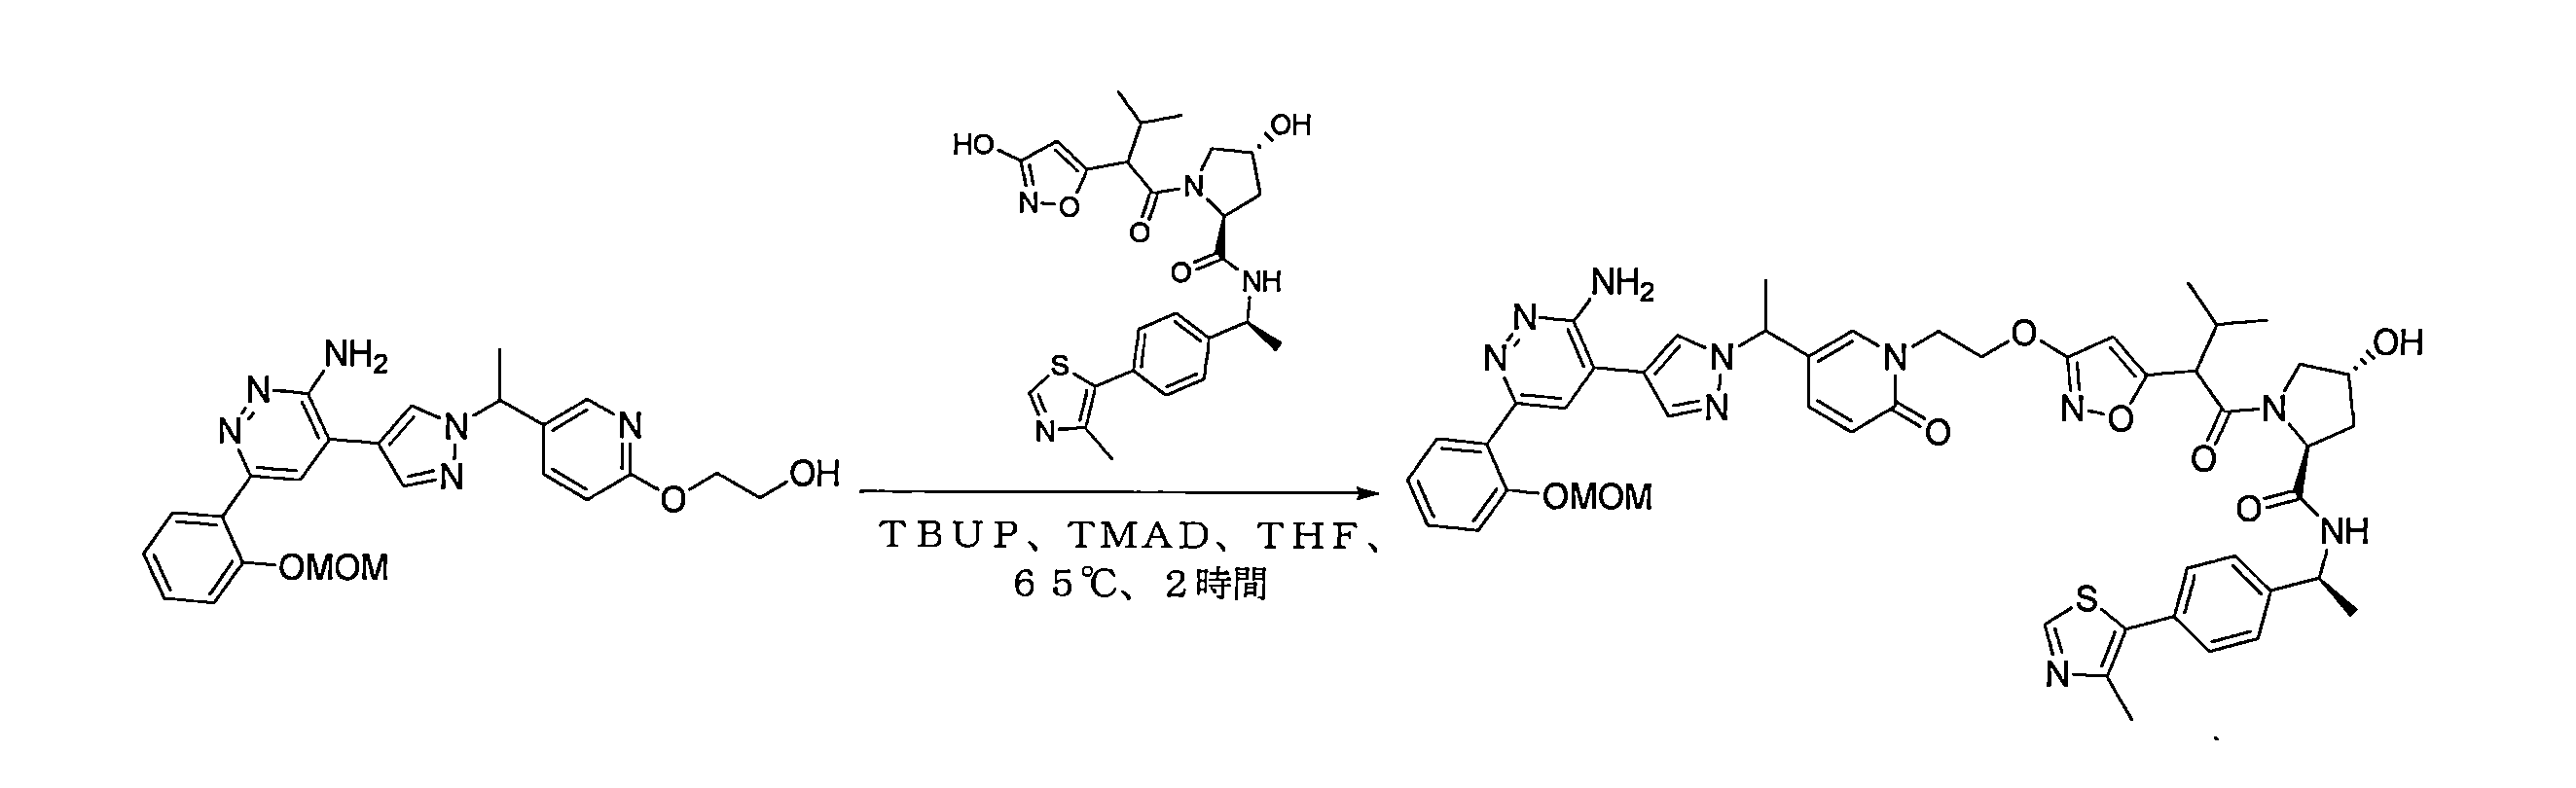
窒素不活性雰囲気でパージし、維持した50mLの丸底フラスコに、THF(15mL)中のテトラメチルアゾジカルボキサミド(861mg、5.00mmol、10.00当量)およびトリブチルホスフィン(1.01g、4.99mmol、9.98当量)の溶液を0℃で入れた。混合物に、2-([5-[1-(4-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-1H-ピラゾール-1-イル)エチル]ピリジン-2-イル]オキシ)エタン-1-オール(231.3mg、0.50mmol、1当量)および(2S,4R)-4-ヒドロキシ-1-[2-(3-ヒドロキシ-1,2-オキサゾール-5-イル)-3-メチルブタノイル]-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(249mg、0.50mmol、1.00当量)の混合物を添加した。得られた溶液を、0℃で10分間撹拌し、次いで、油浴中で、50℃でさらに2時間撹拌した。反応混合物を室温まで冷却し、水(150mL)で希釈した。得られた混合物を酢酸エチル(150mL)で抽出し、有機層をブライン(150mL×2)で洗浄し、無水硫酸ナトリウムで乾燥させ、真空下で濃縮した。残渣を、ジクロロメタン/メタノール(10/1)で溶出するシリカゲルカラム上に適用した。得られた生成物を、以下の条件で分取HPLCによってさらに精製した。カラム、XBridge Prep OBD C18 Column、19×150mm、5um;移動相、水(10mmol/L NH4HCO3を含む)およびアセトニトリル、35% ACN、8分間で最大53%。これにより、68mg(14%)の(2S,4R)-1-(2-(3-(2-(5-(1-(4-(3-アミノ-6-(2-(メトキシメトキシ)フェニル)ピリダジン-4-イル)-1H-ピラゾール-1-イル)エチル)-2-オキソピリジン-1(2H)-イル)エトキシ)イソキサゾール-5-イル)-3-メチルブタノイル)-4-ヒドロキシ-N-((S)-1-(4-(4-メチルチアゾール-5-イル)フェニル)エチル)ピロリジン-2-カルボキサミドを灰色がかった白色の固体として得た。 A 50 mL round bottom flask purged and maintained with a nitrogen inert atmosphere was charged with a solution of tetramethylazodicarboxamide (861 mg, 5.00 mmol, 10.00 equiv.) and tributylphosphine (1.01 g, 4.99 mmol, 9.98 equiv.) in THF (15 mL) at 0°C. To the mixture was added a mixture of 2-([5-[1-(4-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-1H-pyrazol-1-yl)ethyl]pyridin-2-yl]oxy)ethan-1-ol (231.3 mg, 0.50 mmol, 1 equiv.) and (2S,4R)-4-hydroxy-1-[2-(3-hydroxy-1,2-oxazol-5-yl)-3-methylbutanoyl]-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (249 mg, 0.50 mmol, 1.00 equiv.). The resulting solution was stirred at 0° C. for 10 min and then at 50° C. in an oil bath for an additional 2 h. The reaction mixture was cooled to room temperature and diluted with water (150 mL). The resulting mixture was extracted with ethyl acetate (150 mL), and the organic layer was washed with brine (150 mL x 2), dried over anhydrous sodium sulfate, and concentrated under vacuum. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (10/1). The resulting product was further purified by preparative HPLC with the following conditions: Column, XBridge Prep OBD C18 Column, 19 x 150 mm, 5 um; Mobile phase, water (containing 10 mmol/L NH4HCO3) and acetonitrile, 35% ACN, up to 53% in 8 min. This gave 68 mg (14%) of (2S,4R)-1-(2-(3-(2-(5-(1-(4-(3-amino-6-(2-(methoxymethoxy)phenyl)pyridazin-4-yl)-1H-pyrazol-1-yl)ethyl)-2-oxopyridin-1(2H)-yl)ethoxy)isoxazol-5-yl)-3-methylbutanoyl)-4-hydroxy-N-((S)-1-(4-(4-methylthiazol-5-yl)phenyl)ethyl)pyrrolidine-2-carboxamide as an off-white solid.
上記の他の実施例に記載されるものと類似の手順を使用して、(2S,4R)-1-(2-(3-(2-(5-(1-(4-(3-アミノ-6-(2-(メトキシメトキシ)フェニル)ピリダジン-4-イル)-1H-ピラゾール-1-イル)エチル)-2-オキソピリジン-1(2H)-イル)エトキシ)イソキサゾール-5-イル)-3-メチルブタノイル)-4-ヒドロキシ-N-((S)-1-(4-(4-メチルチアゾール-5-イル)フェニル)エチル)ピロリジン-2-カルボキサミドを最終化合物に変換した。 Using procedures similar to those described in other examples above, (2S,4R)-1-(2-(3-(2-(5-(1-(4-(3-amino-6-(2-(methoxymethoxy)phenyl)pyridazin-4-yl)-1H-pyrazol-1-yl)ethyl)-2-oxopyridin-1(2H)-yl)ethoxy)isoxazol-5-yl)-3-methylbutanoyl)-4-hydroxy-N-((S)-1-(4-(4-methylthiazol-5-yl)phenyl)ethyl)pyrrolidine-2-carboxamide was converted to the final compound.
例示的な化合物157を、類似の手順を使用して調製した。 Exemplary compound 157 was prepared using a similar procedure.
例示的な化合物100の例示的な合成
工程1
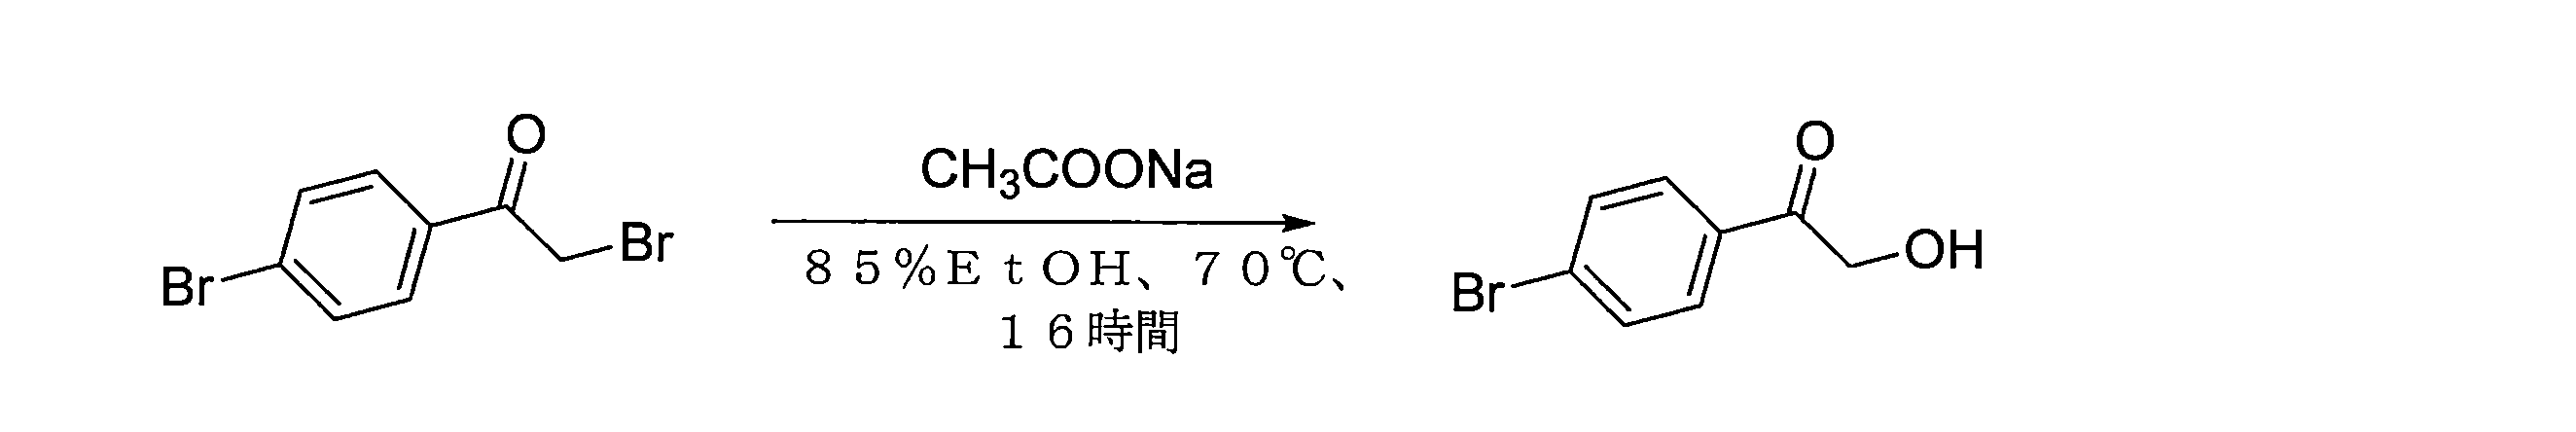
室温のエタノール(350mL、7.60mmol、0.04当量)および水(60mL、3.33mmol、0.02当量)の撹拌混合物に、無水酢酸ナトリウム(18g、264.71mmol、1.47当量)を窒素雰囲気下で少しずつ添加した。得られた混合物を窒素雰囲気下で、70℃で16時間撹拌した。結果として生じる混合物を減圧下で濃縮した。得られた混合物を水(500mL)で希釈した。水層を酢酸エチル(300mL×2)で抽出した。残渣をブライン(100mL×2)で洗浄し、乾燥させ、減圧下で濃縮した。残渣を、酢酸エチル/石油エーテル(1:5)で溶出するシリカゲルカラムクロマトグラフィーによって精製して、1-(4-ブロモフェニル)-2-ヒドロキシエタン-1-オン(32g、83%)を白色の固体として得た。 To a stirred mixture of ethanol (350 mL, 7.60 mmol, 0.04 equiv.) and water (60 mL, 3.33 mmol, 0.02 equiv.) at room temperature, anhydrous sodium acetate (18 g, 264.71 mmol, 1.47 equiv.) was added portionwise under nitrogen atmosphere. The resulting mixture was stirred at 70° C. for 16 h under nitrogen atmosphere. The resulting mixture was concentrated under reduced pressure. The resulting mixture was diluted with water (500 mL). The aqueous layer was extracted with ethyl acetate (300 mL×2). The residue was washed with brine (100 mL×2), dried and concentrated under reduced pressure. The residue was purified by silica gel column chromatography eluting with ethyl acetate/petroleum ether (1:5) to give 1-(4-bromophenyl)-2-hydroxyethan-1-one (32 g, 83%) as a white solid.
工程2

1Lの丸底フラスコに、1-(4-ブロモフェニル)-2-ヒドロキシエタン-1-オン(32g、148.8mmol、1当量)、塩化メチレン(500mL)、イミダゾール(30g、440.7mmol、2.96当量)、4-ジメチルアミノピリジン(1.8g、14.73mmol、0.10当量)、およびtert-ブチルジメチルシリルクロリド(26.8g、177.8mmol、1.19当量)を窒素雰囲気下で、室温で添加した。得られた混合物を窒素雰囲気下で、室温で2時間撹拌した。反応物を室温で、水でクエンチした。得られた混合物を塩化メチレン(200mL×3)で抽出した。合わせた有機層をブライン(100mL×2)で洗浄し、無水硫酸ナトリウムで乾燥させ、減圧下で濃縮した。残渣を、酢酸エチル/石油エーテル(1:20)で溶出するシリカゲルカラムクロマトグラフィーによって精製して、1-(4-ブロモフェニル)-2-[(tert-ブチルジメチルシリル)オキシ]エタン-1-オン(29g、59%)を淡い黄色の固体として得た。 In a 1 L round bottom flask, 1-(4-bromophenyl)-2-hydroxyethan-1-one (32 g, 148.8 mmol, 1 equiv.), methylene chloride (500 mL), imidazole (30 g, 440.7 mmol, 2.96 equiv.), 4-dimethylaminopyridine (1.8 g, 14.73 mmol, 0.10 equiv.), and tert-butyldimethylsilyl chloride (26.8 g, 177.8 mmol, 1.19 equiv.) were added at room temperature under nitrogen atmosphere. The resulting mixture was stirred at room temperature under nitrogen atmosphere for 2 hours. The reaction was quenched with water at room temperature. The resulting mixture was extracted with methylene chloride (200 mL x 3). The combined organic layers were washed with brine (100 mL x 2), dried over anhydrous sodium sulfate, and concentrated under reduced pressure. The residue was purified by silica gel column chromatography eluting with ethyl acetate/petroleum ether (1:20) to give 1-(4-bromophenyl)-2-[(tert-butyldimethylsilyl)oxy]ethan-1-one (29 g, 59%) as a pale yellow solid.
工程3

1Lの三口丸底フラスコに、1-(4-ブロモフェニル)-2-[(tert-ブチルジメチルシリル)オキシ]エタン-1-オン(27g、81.99mmol、1当量)、テトラヒドロフラン(500mL)、および(S)-2-メチルプロパン-2-スルフィンアミド(19.5g、160.9mmol、1.96当量)を室温で添加した。撹拌溶液に、Ti(i-PrO)4(70g、246.5mmol、3.01当量)を窒素雰囲気下で、室温で、滴下で添加した。得られた混合物を窒素雰囲気下で、70℃で16時間撹拌した。混合物を室温まで冷却させた。得られた混合物を濾過し、フィルターケーキを酢酸エチル(200mL×2)で洗浄した。合わせた有機層をブライン(200mL)で洗浄し、硫酸ナトリウムで乾燥させ、減圧下で濃縮した。残渣を、酢酸エチル/石油エーテル(1:15)で溶出するシリカゲルカラムクロマトグラフィーによって精製して、(S)-N-[(1Z)-1-(4-ブロモフェニル)-2-[(tert-ブチルジメチルシリル)オキシ]エチリデン]-2-メチルプロパン-2-スルフィンアミド(16g、45%)を黄色の油として得た。 In a 1 L three-neck round bottom flask, 1-(4-bromophenyl)-2-[(tert-butyldimethylsilyl)oxy]ethan-1-one (27 g, 81.99 mmol, 1 eq.), tetrahydrofuran (500 mL), and (S)-2-methylpropane-2-sulfinamide (19.5 g, 160.9 mmol, 1.96 eq.) were added at room temperature. To the stirred solution, Ti(i-PrO)4 (70 g, 246.5 mmol, 3.01 eq.) was added dropwise at room temperature under nitrogen atmosphere. The resulting mixture was stirred at 70° C. for 16 hours under nitrogen atmosphere. The mixture was allowed to cool to room temperature. The resulting mixture was filtered and the filter cake was washed with ethyl acetate (200 mL×2). The combined organic layers were washed with brine (200 mL), dried over sodium sulfate, and concentrated under reduced pressure. The residue was purified by silica gel column chromatography eluting with ethyl acetate/petroleum ether (1:15) to give (S)-N-[(1Z)-1-(4-bromophenyl)-2-[(tert-butyldimethylsilyl)oxy]ethylidene]-2-methylpropane-2-sulfinamide (16 g, 45%) as a yellow oil.
工程4

500mLの三口丸底フラスコに、(S)-N-[(1Z)-1-(4-ブロモフェニル)-2-[(tert-ブチルジメチルシリル)オキシ]エチリデン]-2-メチルプロパン-2-スルフィンアミド(15g、34.68mmol、1当量)およびテトラヒドロフラン(150mL)を室温で添加した。撹拌溶液に、BH3.THF(52mL、52mmol、1.5当量)を窒素雰囲気下で、-70℃で、滴下で添加した。得られた混合物を、窒素雰囲気下で、-70℃で1時間撹拌した。反応物を室温で、メタノールでクエンチした。得られた混合物を濾過し、フィルターケーキを酢酸エチル(100mL×2)で洗浄した。濾液をブライン(50mL×2)で洗浄し、無水硫酸ナトリウムで乾燥させ、減圧下で濃縮した。残渣を、酢酸エチル/石油エーテル(1:2)で溶出するシリカゲルカラムクロマトグラフィーによって精製して、(S)-N-[(1R)-1-(4-ブロモフェニル)-2-[(tert-ブチルジメチルシリル)オキシ]エチル]-2-メチルプロパン-2-スルフィンアミド(5.6g、37%)を黄色の固体として得た。 In a 500 mL three-neck round bottom flask, (S)-N-[(1Z)-1-(4-bromophenyl)-2-[(tert-butyldimethylsilyl)oxy]ethylidene]-2-methylpropane-2-sulfinamide (15 g, 34.68 mmol, 1 eq.) and tetrahydrofuran (150 mL) were added at room temperature. To the stirred solution, BH3.THF (52 mL, 52 mmol, 1.5 eq.) was added dropwise at −70° C. under nitrogen atmosphere. The resulting mixture was stirred at −70° C. for 1 h under nitrogen atmosphere. The reaction was quenched with methanol at room temperature. The resulting mixture was filtered and the filter cake was washed with ethyl acetate (100 mL×2). The filtrate was washed with brine (50 mL×2), dried over anhydrous sodium sulfate, and concentrated under reduced pressure. The residue was purified by silica gel column chromatography eluting with ethyl acetate/petroleum ether (1:2) to give (S)-N-[(1R)-1-(4-bromophenyl)-2-[(tert-butyldimethylsilyl)oxy]ethyl]-2-methylpropane-2-sulfinamide (5.6 g, 37%) as a yellow solid.
工程5
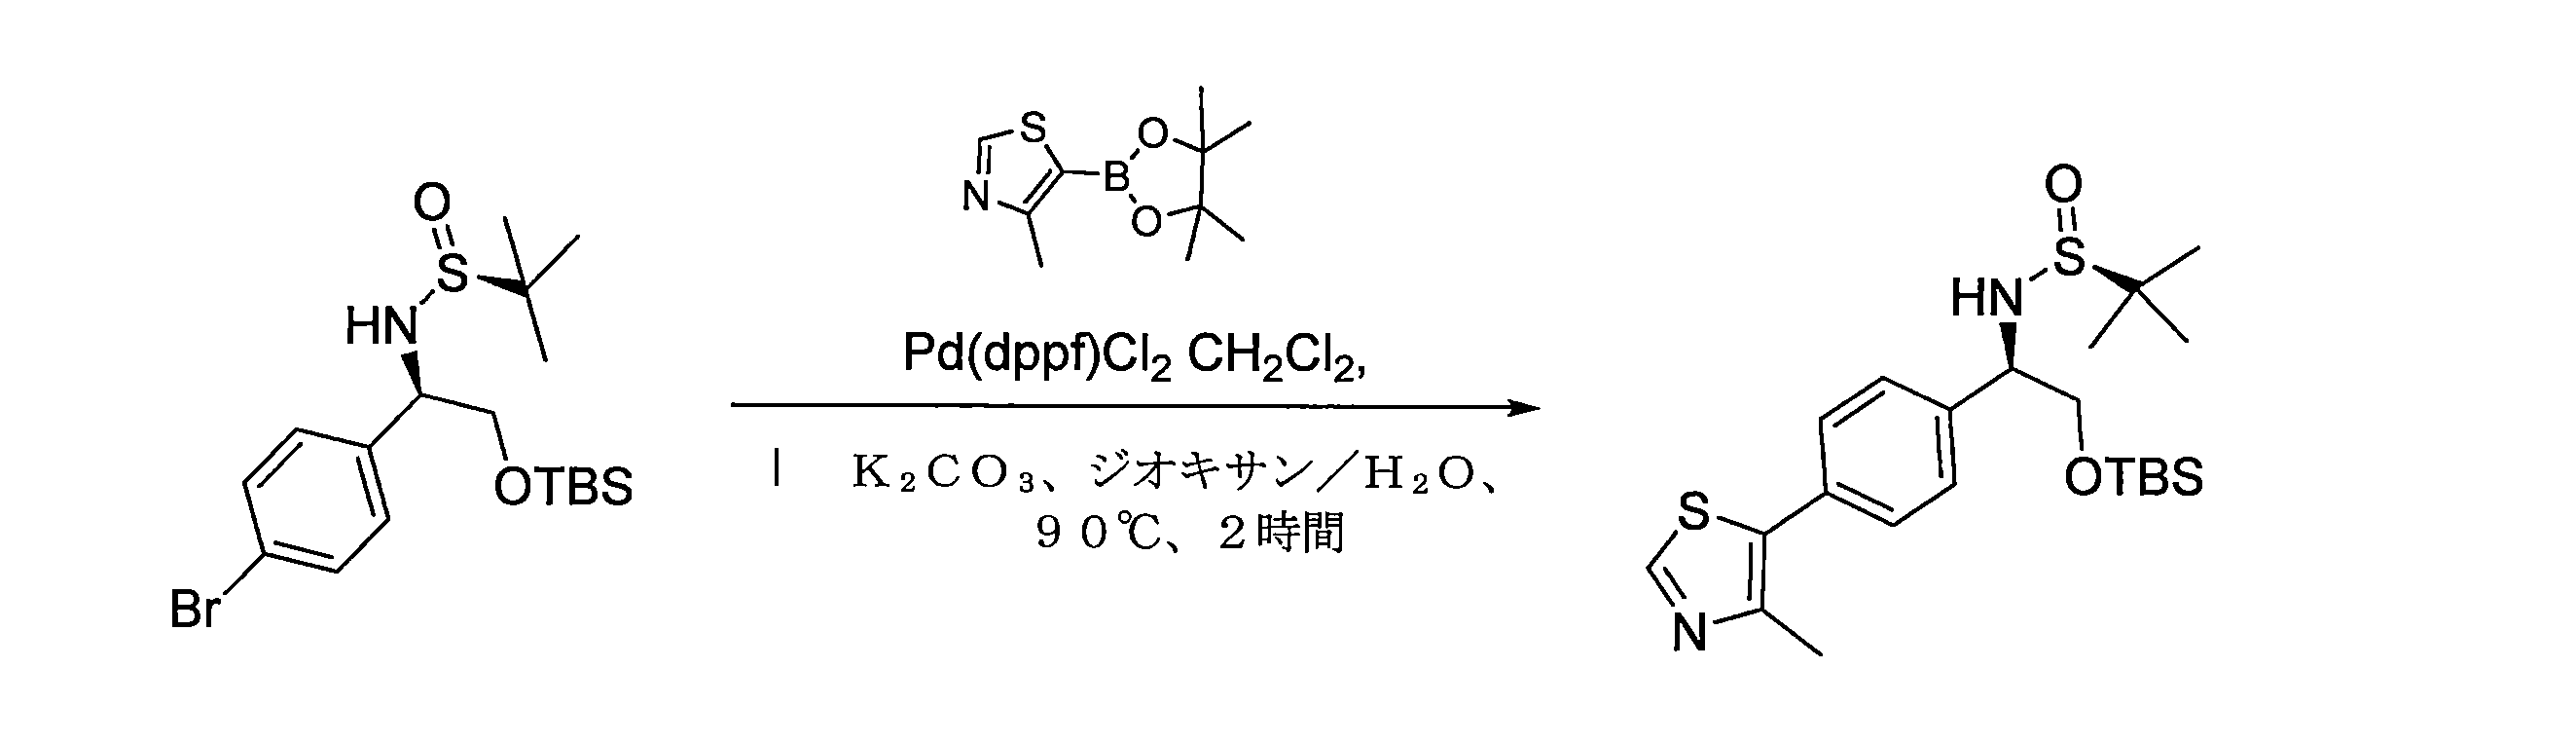
窒素不活性雰囲気下でパージし、維持した100mLの丸底フラスコに、(S)-N-[(1R)-1-(4-ブロモフェニル)-2-[(tert-ブチルジメチルシリル)オキシ]エチル]-2-メチルプロパン-2-スルフィンアミド(2.6g、5.98mmol、1.00当量)、ジオキサン(30mL)、4-メチル-5-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)-1,3-チアゾール(2g、8.88mmol、1.48当量)、炭酸カリウム(2.5g、18.09mmol、3.02当量)、水(5mL)、およびPd(dppf)Cl2.CH2Cl2(440mg、0.51mmol、0.09当量)を入れた。得られた溶液を90℃で2時間撹拌した。固体を濾別した。濾液を水(50mL)で希釈し、酢酸エチル(50mL×3)で抽出した。合わせた有機層をブライン(35mL×2)で洗浄し、無水硫酸ナトリウムで乾燥させ、真空下で濃縮した。残渣を、酢酸エチル/石油エーテル(1:5)で溶出するシリカゲルカラム上に適用した。これにより、2.5g(92%)の(S)-N-[(1R)-2-[(tert-ブチルジメチルシリル)オキシ]-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]-2-メチルプロパン-2-スルフィンアミドをオレンジ色の油として得た。 A 100 mL round bottom flask purged and maintained under nitrogen inert atmosphere was charged with (S)-N-[(1R)-1-(4-bromophenyl)-2-[(tert-butyldimethylsilyl)oxy]ethyl]-2-methylpropane-2-sulfinamide (2.6 g, 5.98 mmol, 1.00 equiv.), dioxane (30 mL), 4-methyl-5-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)-1,3-thiazole (2 g, 8.88 mmol, 1.48 equiv.), potassium carbonate (2.5 g, 18.09 mmol, 3.02 equiv.), water (5 mL), and Pd(dppf)Cl2.CH2Cl2 (440 mg, 0.51 mmol, 0.09 equiv.). The resulting solution was stirred at 90 °C for 2 h. The solids were filtered off. The filtrate was diluted with water (50 mL) and extracted with ethyl acetate (50 mL x 3). The combined organic layers were washed with brine (35 mL x 2), dried over anhydrous sodium sulfate, and concentrated under vacuum. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (1:5). This gave 2.5 g (92%) of (S)-N-[(1R)-2-[(tert-butyldimethylsilyl)oxy]-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]-2-methylpropane-2-sulfinamide as an orange oil.
工程6
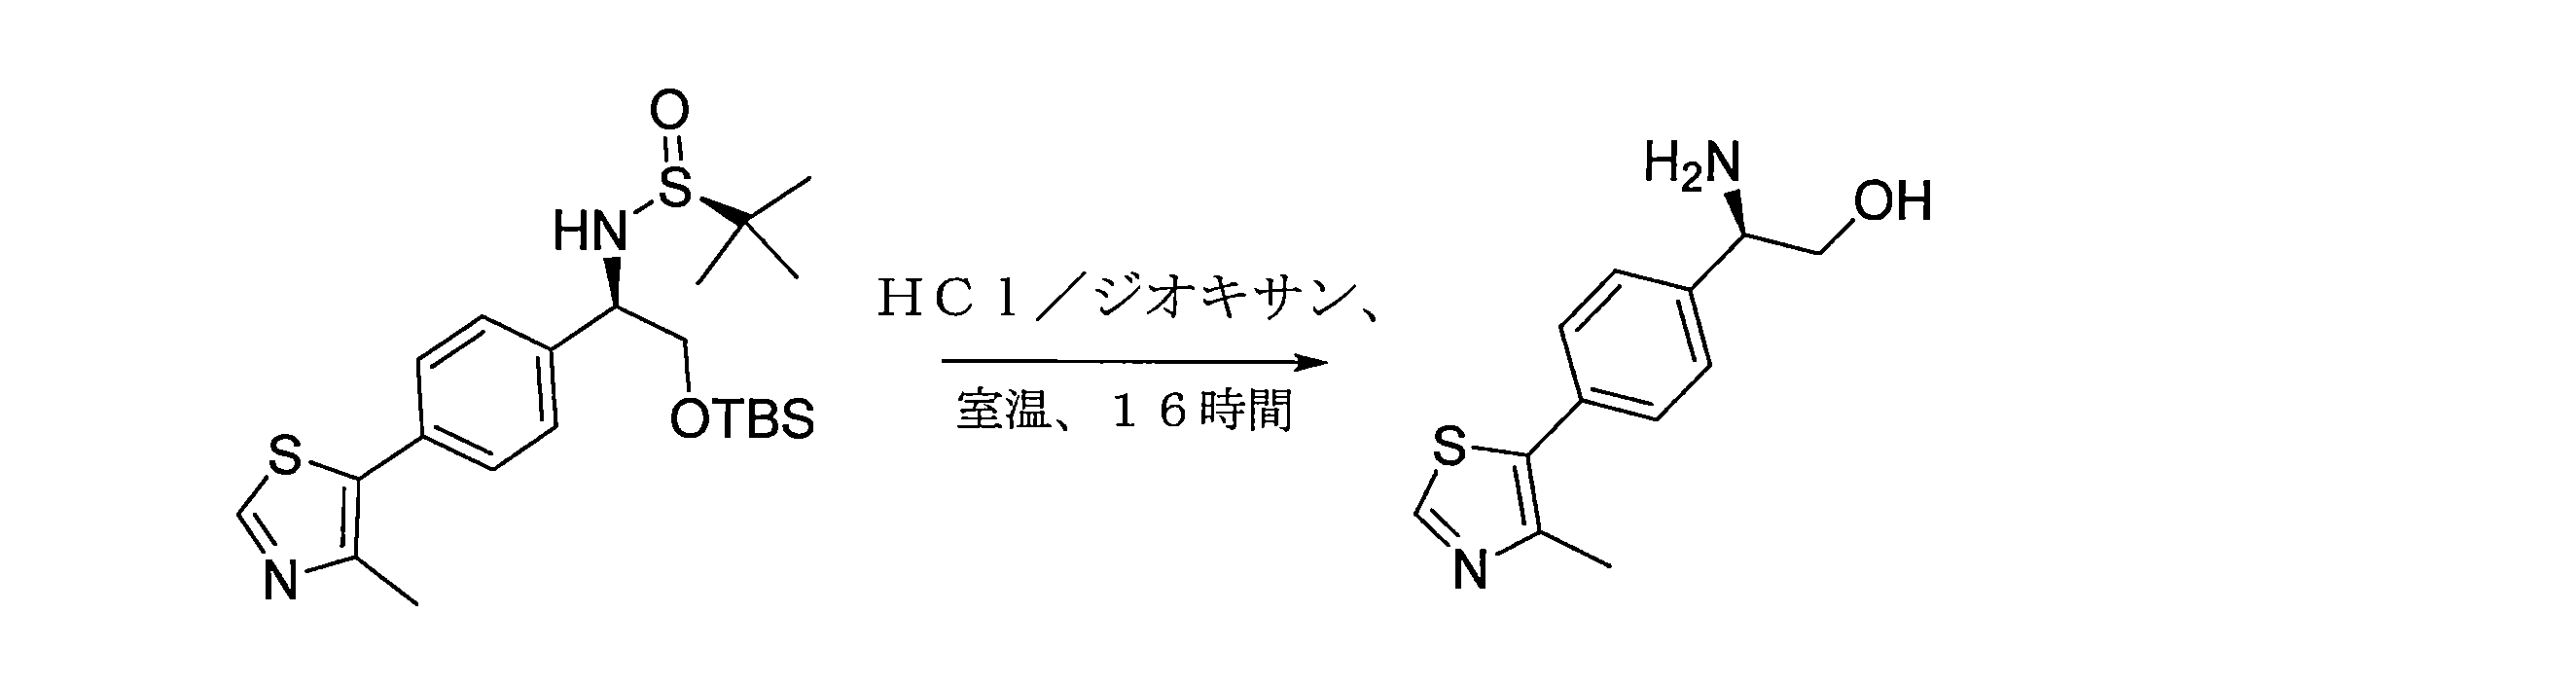
250mLの丸底フラスコに、ジオキサン(100mL、4M)中の(S)-N-[(1R)-2-[(tert-ブチルジメチルシリル)オキシ]-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]-2-メチルプロパン-2-スルフィンアミド(2.6g、5.74mmol、1.00当量)および塩化水素を入れた。得られた溶液を、室温で一晩撹拌した。得られた混合物を真空下で濃縮した。これにより、1.5g(96%)の(2R)-2-アミノ-2-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エタン-1-オール塩酸塩を黄色の油として得た。 A 250 mL round bottom flask was charged with (S)-N-[(1R)-2-[(tert-butyldimethylsilyl)oxy]-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]-2-methylpropane-2-sulfinamide (2.6 g, 5.74 mmol, 1.00 equiv.) in dioxane (100 mL, 4 M) and hydrogen chloride. The resulting solution was stirred overnight at room temperature. The resulting mixture was concentrated under vacuum. This gave 1.5 g (96%) of (2R)-2-amino-2-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethan-1-ol hydrochloride as a yellow oil.
工程7
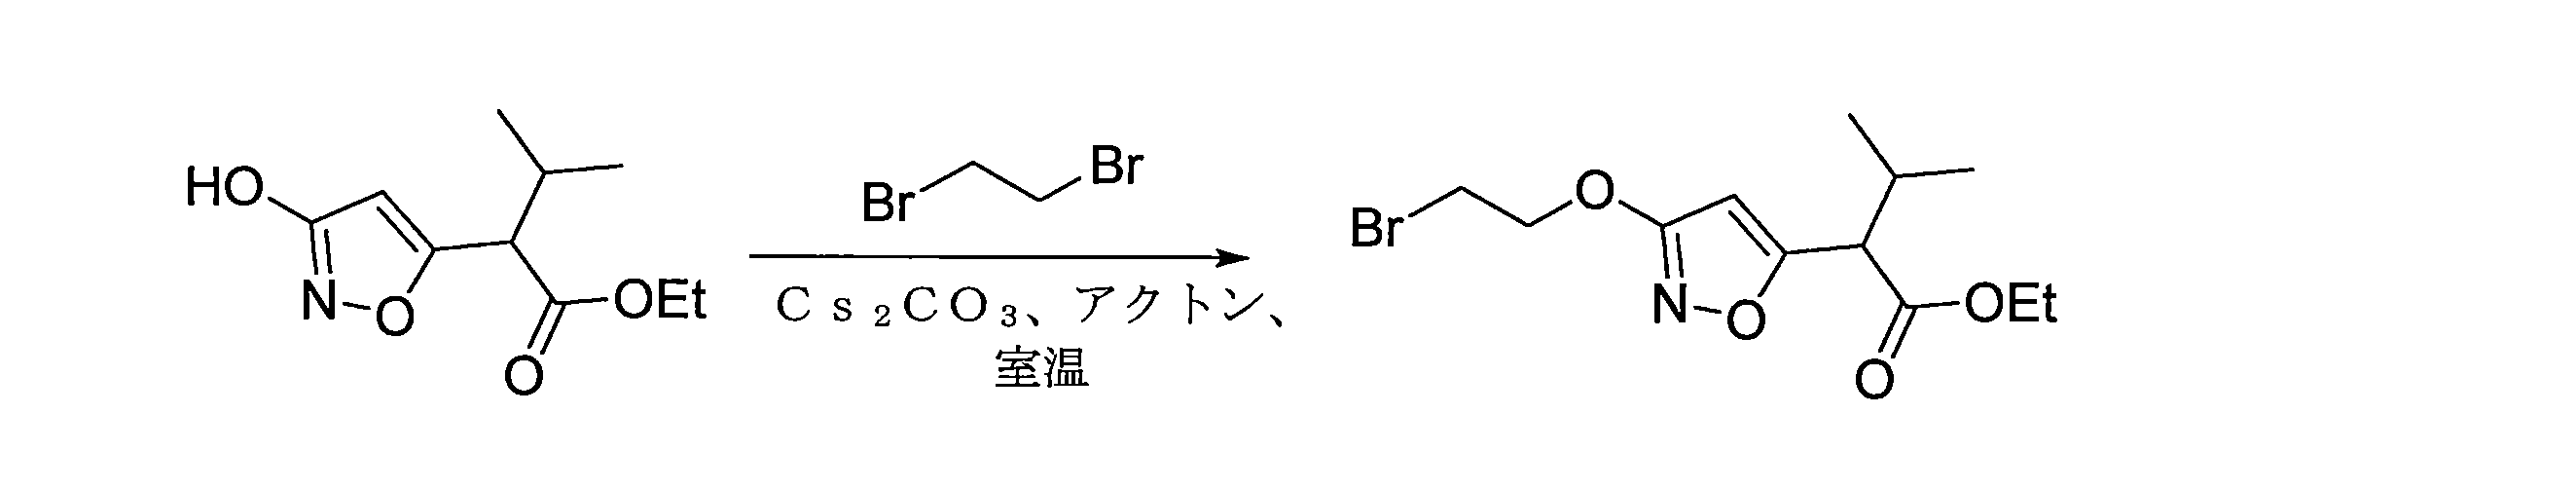
250mLの丸底フラスコに、アセトン(100mL)中のエチル2-(3-ヒドロキシ-1,2-オキサゾール-5-イル)-3-メチルブタノエート(1g、4.69mmol、1当量)、Cs2CO3(4.5g、13.81mmol、3.0当量)、および1,2-ジブロモエタン(2.60g、13.84mmol)を入れた。得られた混合物を室温で一晩撹拌した。次いで、反応物を水(100mL)によってクエンチした。得られた混合物を酢酸エチル(100mL×3)で抽出し、有機層を合わせ、無水硫酸ナトリウムで乾燥させ、減圧下で濃縮した。残渣を、酢酸エチル/石油エーテル(3:10)で溶出するシリカゲルカラム上に適用した。これにより、600mg(40%)のエチル2-[3-(2-ブロモエトキシ)-1,2-オキサゾール-5-イル]-3-メチルブタノエートを淡い黄色の油として得た。 A 250 mL round bottom flask was charged with ethyl 2-(3-hydroxy-1,2-oxazol-5-yl)-3-methylbutanoate (1 g, 4.69 mmol, 1 equiv.), Cs2CO3 (4.5 g, 13.81 mmol, 3.0 equiv.), and 1,2-dibromoethane (2.60 g, 13.84 mmol) in acetone (100 mL). The resulting mixture was stirred at room temperature overnight. The reaction was then quenched by water (100 mL). The resulting mixture was extracted with ethyl acetate (100 mL x 3), the organic layers were combined, dried over anhydrous sodium sulfate, and concentrated under reduced pressure. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (3:10). This afforded 600 mg (40%) of ethyl 2-[3-(2-bromoethoxy)-1,2-oxazol-5-yl]-3-methylbutanoate as a pale yellow oil.
他の実施例について記載され、当業者に明らかである手順を使用して、以下のスキームに従って、(2R)-2-アミノ-2-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エタン-1-オールおよびエチル2-[3-(2-ブロモエトキシ)-1,2-オキサゾール-5-イル]-3-メチルブタノエートを、(2S,4R)-1-(2-(3-(2-ブロモエトキシ)イソキサゾール-5-イル)-3-メチルブタノイル)-4-ヒドロキシ-N-((R)-2-ヒドロキシ-1-(4-(4-メチルチアゾール-5-イル)フェニル)エチル)ピロリジン-2-カルボキサミドに変換した。
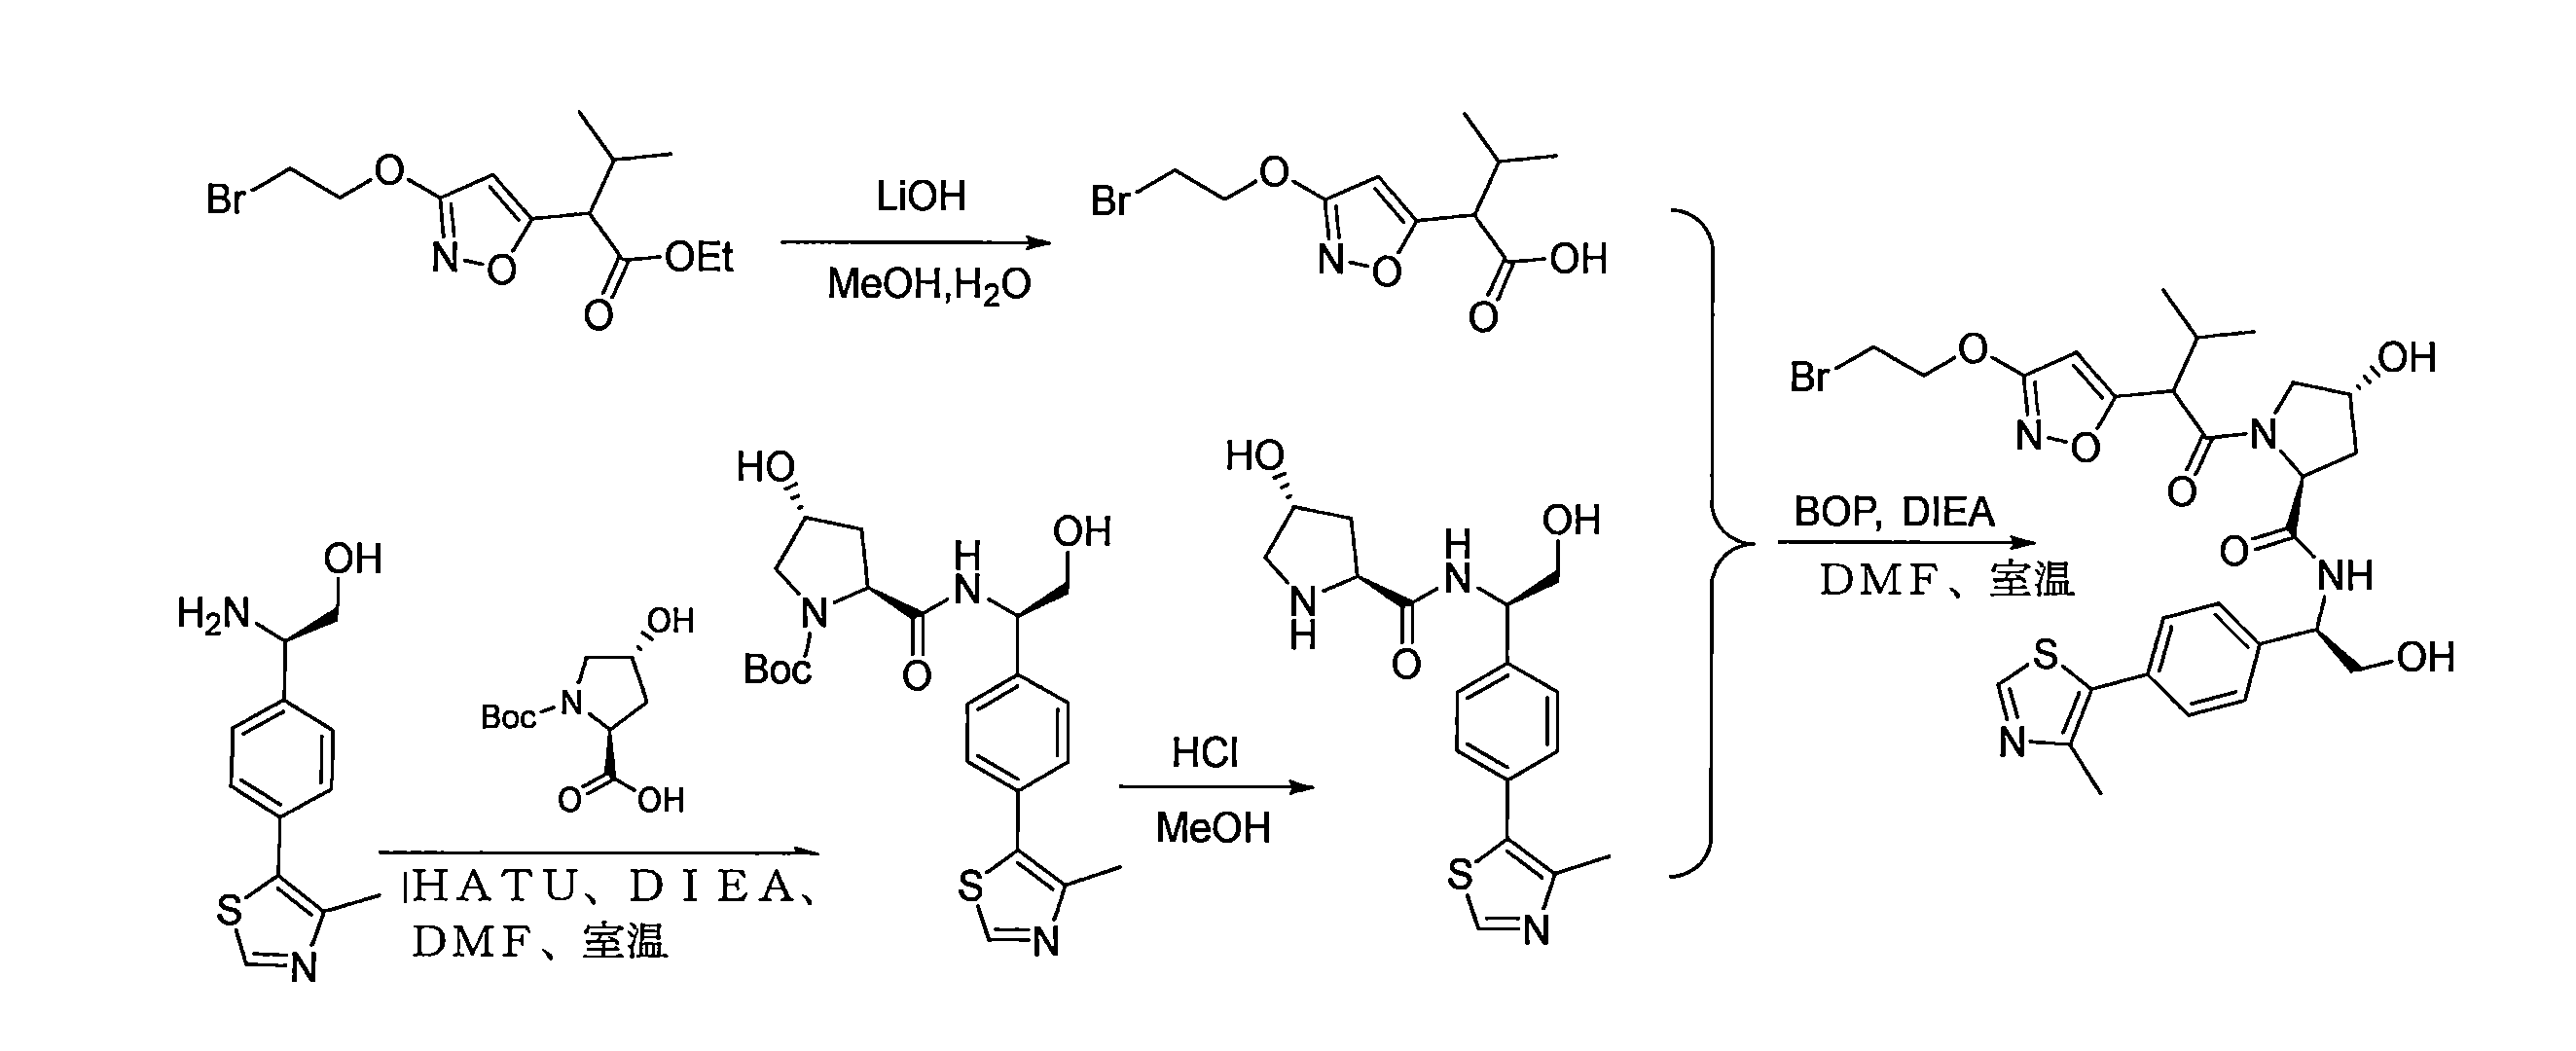
例えば、例示的な化合物47に記載されるものと類似の手順を使用して、(2S,4R)-1-(2-(3-(2-ブロモエトキシ)イソキサゾール-5-イル)-3-メチルブタノイル)-4-ヒドロキシ-N-((R)-2-ヒドロキシ-1-(4-(4-メチルチアゾール-5-イル)フェニル)エチル)ピロリジン-2-カルボキサミドを最終化合物に変換した。 For example, (2S,4R)-1-(2-(3-(2-bromoethoxy)isoxazol-5-yl)-3-methylbutanoyl)-4-hydroxy-N-((R)-2-hydroxy-1-(4-(4-methylthiazol-5-yl)phenyl)ethyl)pyrrolidine-2-carboxamide was converted to the final compound using a procedure similar to that described in exemplary compound 47.
例示的な化合物158を、類似の手順を使用して調製した。 Exemplary compound 158 was prepared using a similar procedure.
例示的な化合物115の例示的な合成
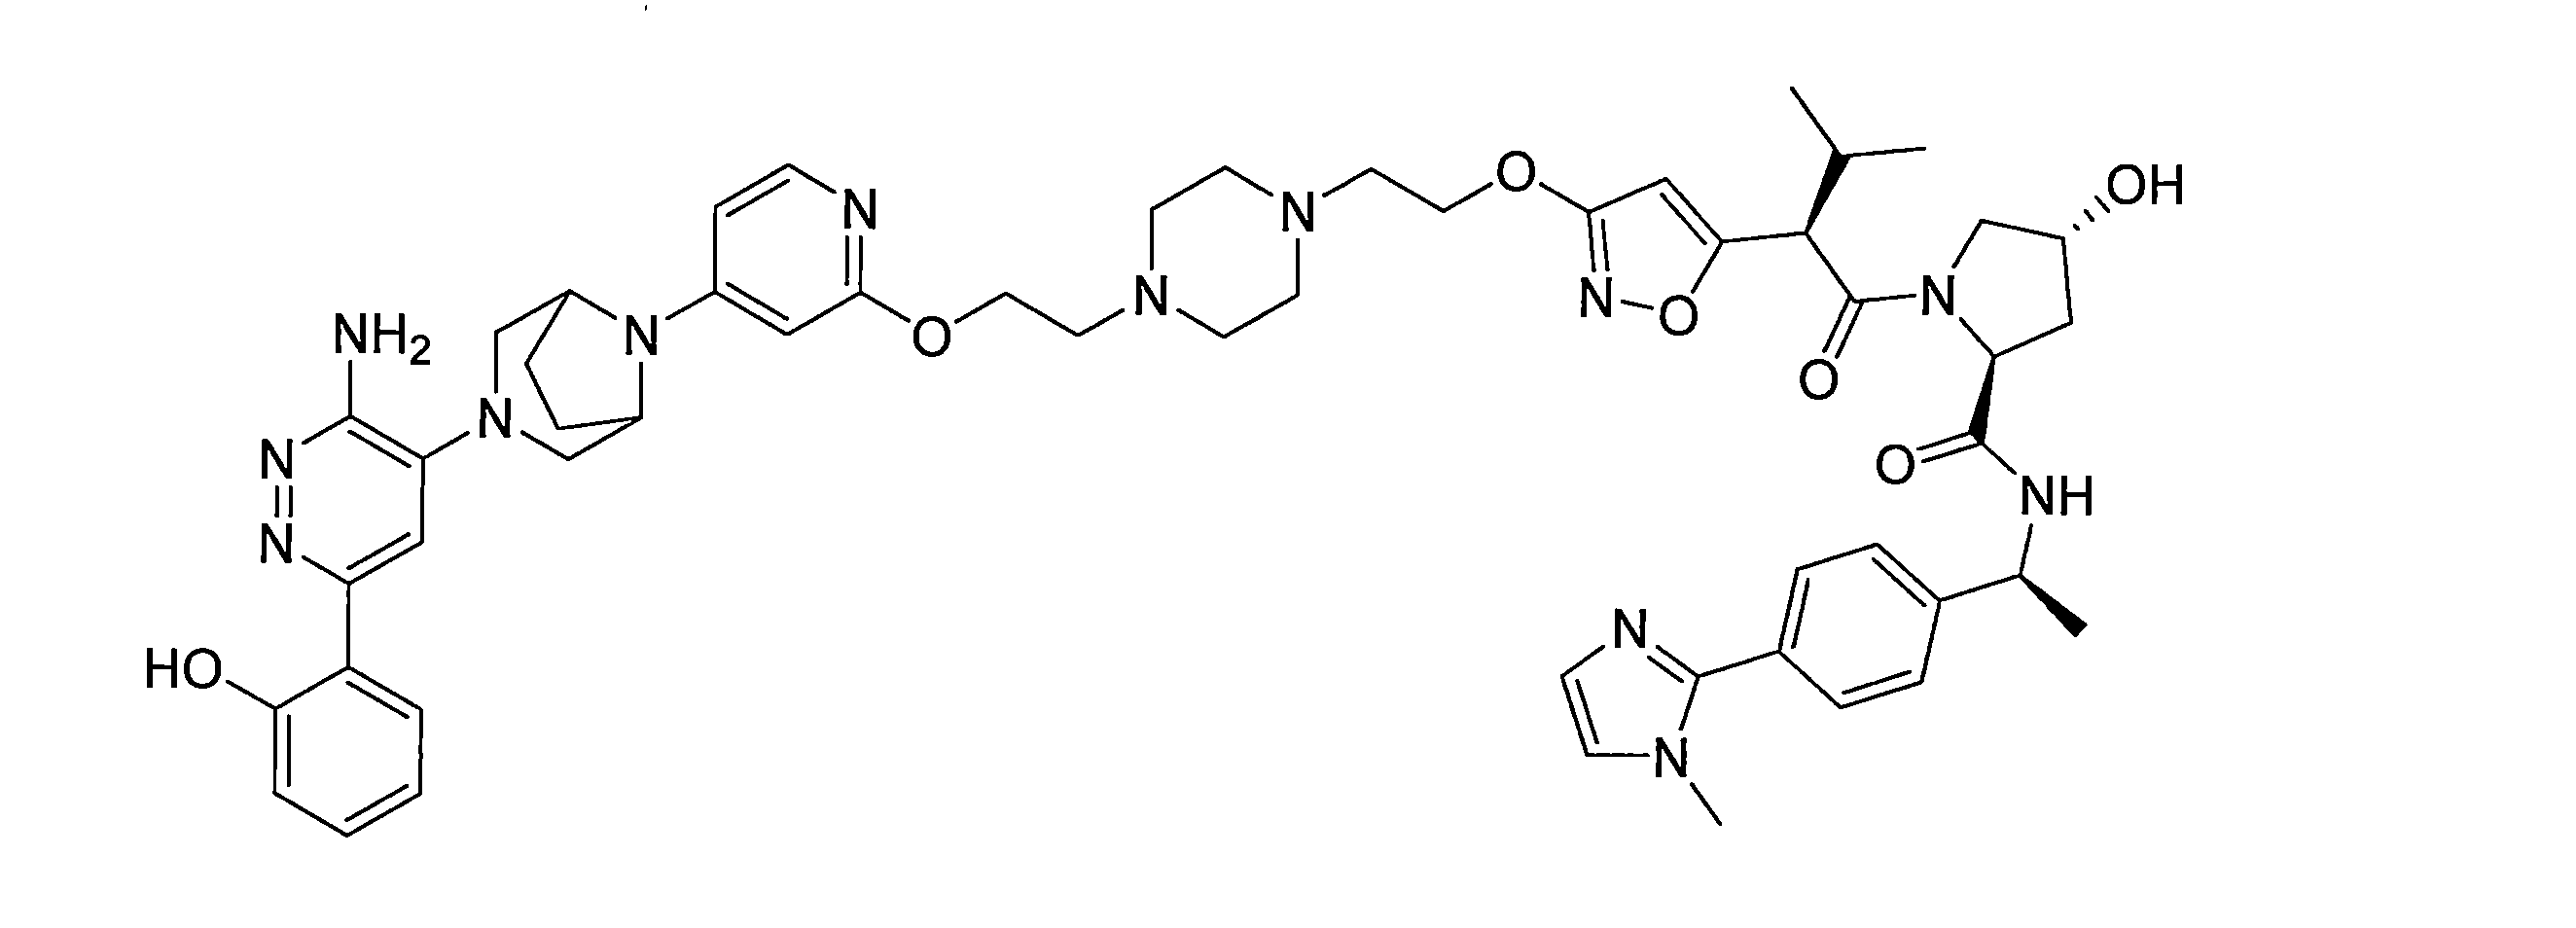
工程1
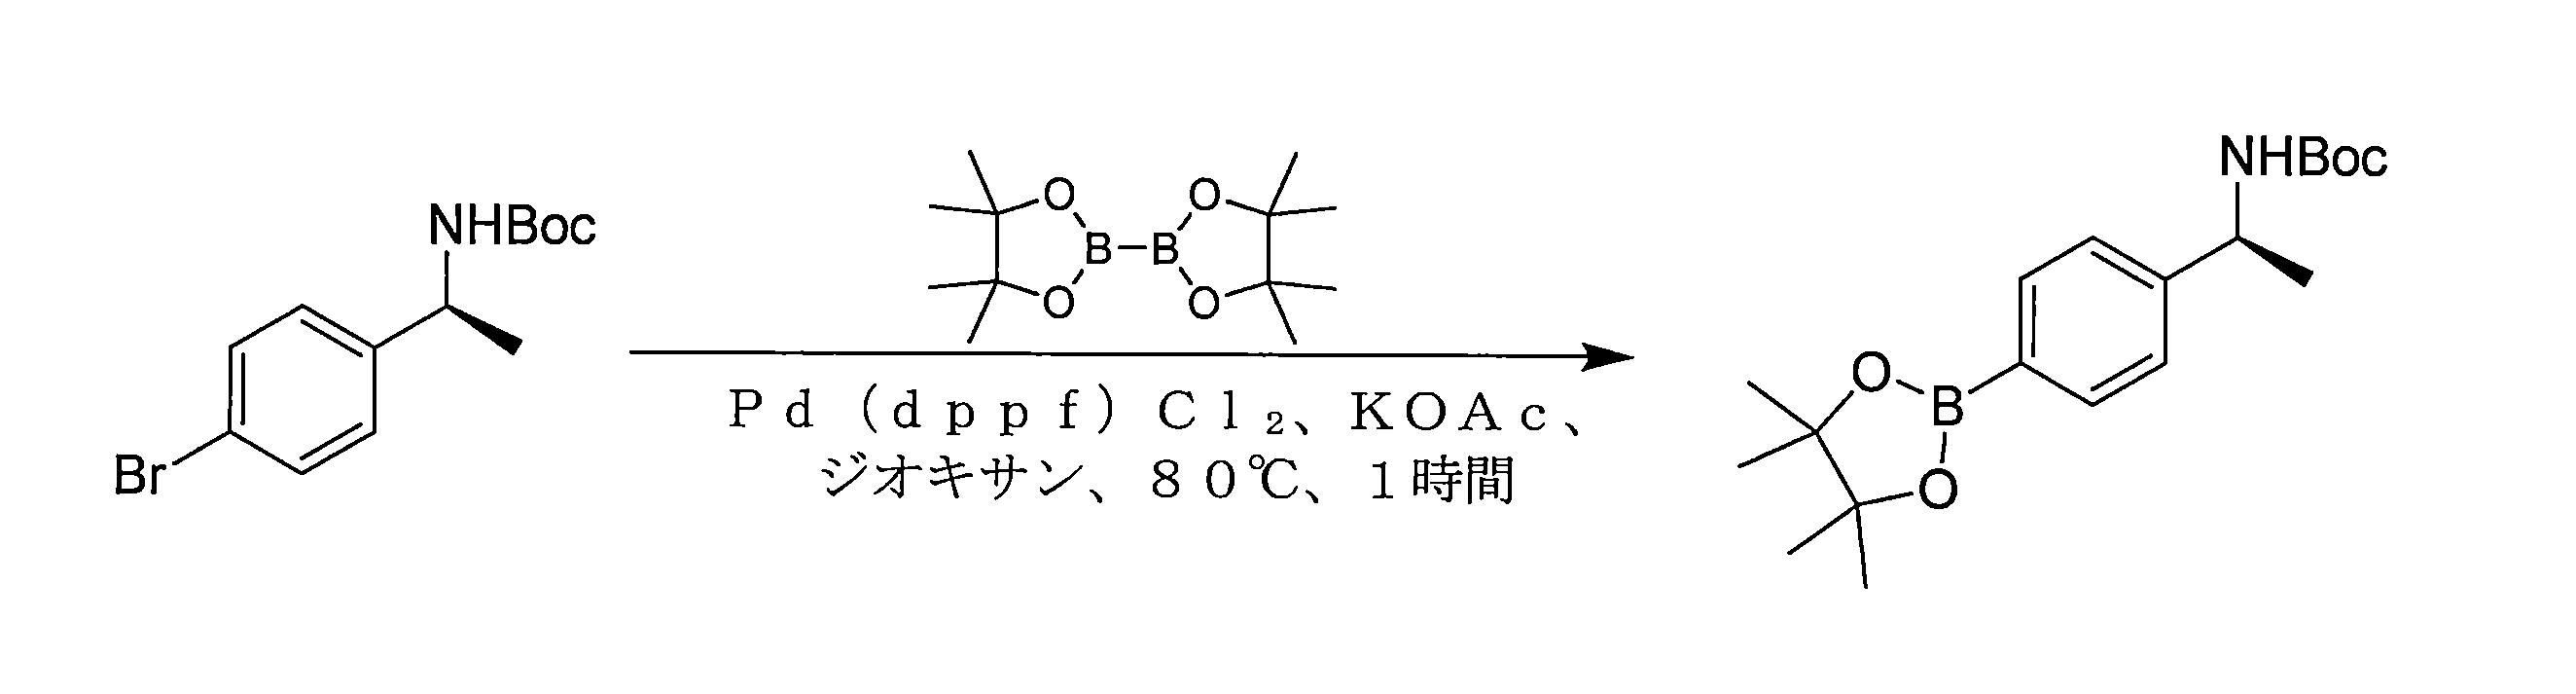
フラスコに、tert-ブチルN-[(1S)-1-(4-ブロモフェニル)エチル]カルバメート(1.4g、4.66mmol、1当量)、4,4,5,5-テトラメチル-2-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)-1,3,2-ジオキサボロラン(1.42g、5.60mmol、1.2当量)、(1,1’-ビス(ジフェニルホスフィノ)フェロセン)パラジウム(II)二塩化物(170mg、0.23mmol、0.05当量)、酢酸カリウム(915mg、9.33mmol、2当量)、およびジオキサン(30mL)を充填した。混合物を窒素で10分間パージし、次いで、80℃に1時間加熱した。反応混合物を20℃まで冷却し、セライトのパッドを通して濾過した。濾液を真空中で濃縮した。粗生成物をカラム(石油エーテル:酢酸エチル=10:1)によって精製した。Tert-ブチルN-[(1S)-1-[4-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)フェニル]エチル]カルバメート(1.9g、粗製)を、無色の油として得た。 A flask was charged with tert-butyl N-[(1S)-1-(4-bromophenyl)ethyl]carbamate (1.4 g, 4.66 mmol, 1 equiv.), 4,4,5,5-tetramethyl-2-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)-1,3,2-dioxaborolane (1.42 g, 5.60 mmol, 1.2 equiv.), (1,1'-bis(diphenylphosphino)ferrocene)palladium(II) dichloride (170 mg, 0.23 mmol, 0.05 equiv.), potassium acetate (915 mg, 9.33 mmol, 2 equiv.), and dioxane (30 mL). The mixture was purged with nitrogen for 10 min and then heated to 80 °C for 1 h. The reaction mixture was cooled to 20 °C and filtered through a pad of Celite. The filtrate was concentrated in vacuo. The crude product was purified by column (petroleum ether: ethyl acetate = 10:1). Tert-butyl N-[(1S)-1-[4-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)phenyl]ethyl]carbamate (1.9 g, crude) was obtained as a colorless oil.
工程2

フラスコに、tert-ブチルN-[(1S)-1-[4-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)フェニル]エチル]カルバメート(1.90g、5.47mmol、1.1当量)、2-ブロモ-1-メチル-イミダゾール(800mg、4.97mmol、1当量)、(1,1’-ビス(ジフェニルホスフィノ)フェロセン)パラジウム(II)二塩化物(181mg、0.24mmol、0.05当量)、炭酸ナトリウム(1.05g、9.94mmol、2当量)、ジオキサン(18mL)、および水(3mL)を充填した。混合物を窒素で5分間パージし、110℃に12時間加熱した。混合物を水(50mL)で希釈し、酢酸エチル(20mL×3)で抽出した。有機層をブライン(50mL)で洗浄し、硫酸ナトリウムで乾燥させ、濾過した。濾液を真空中で濃縮した。粗残渣をカラムクロマトグラフィー(石油エーテル:酢酸エチル=1:0~3:1)によって精製した。Tert-ブチルN-[(1S)-1-[4-(1-メチルイミダゾール-2-イル)フェニル]エチル]カルバメート(850mg、2.82mmol、収率56%)を灰色の固体として得た。 A flask was charged with tert-butyl N-[(1S)-1-[4-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)phenyl]ethyl]carbamate (1.90 g, 5.47 mmol, 1.1 eq.), 2-bromo-1-methyl-imidazole (800 mg, 4.97 mmol, 1 eq.), (1,1'-bis(diphenylphosphino)ferrocene)palladium(II) dichloride (181 mg, 0.24 mmol, 0.05 eq.), sodium carbonate (1.05 g, 9.94 mmol, 2 eq.), dioxane (18 mL), and water (3 mL). The mixture was purged with nitrogen for 5 minutes and heated to 110 °C for 12 hours. The mixture was diluted with water (50 mL) and extracted with ethyl acetate (20 mL x 3). The organic layer was washed with brine (50 mL), dried over sodium sulfate, and filtered. The filtrate was concentrated in vacuo. The crude residue was purified by column chromatography (petroleum ether:ethyl acetate=1:0 to 3:1). Tert-butyl N-[(1S)-1-[4-(1-methylimidazol-2-yl)phenyl]ethyl]carbamate (850 mg, 2.82 mmol, 56% yield) was obtained as a grey solid.
工程3

ジオキサン(2mL)中のtert-ブチルN-[(1S)-1-[4-(1-メチルイミダゾール-2-イル)フェニル]エチル]カルバメート(850mg、2.82mmol、1当量)の溶液に、塩化水素溶液(ジオキサン中4M、10mL、14.18当量)を添加した。溶液を20℃で1時間撹拌した。溶媒を真空中で除去した。(1S)-1-[4-(1-メチルイミダゾール-2-イル)フェニル]エタンアミン(670mg、粗製、塩酸塩)を灰色の固体として得た。 To a solution of tert-butyl N-[(1S)-1-[4-(1-methylimidazol-2-yl)phenyl]ethyl]carbamate (850 mg, 2.82 mmol, 1 equiv.) in dioxane (2 mL) was added hydrogen chloride solution (4 M in dioxane, 10 mL, 14.18 equiv.). The solution was stirred at 20°C for 1 h. The solvent was removed in vacuo. (1S)-1-[4-(1-methylimidazol-2-yl)phenyl]ethanamine (670 mg, crude, hydrochloride salt) was obtained as a grey solid.
工程4

N,N-ジメチルホルムアミド(2mL)中の(2S,4R)-1-tert-ブトキシカルボニル-4-ヒドロキシ-ピロリジン-2-カルボン酸(651mg、2.82mmol、1当量)の溶液に、1-ヒドロキシベンゾトリアゾール(456mg、3.38mmol、1.2当量)、1-(3-ジメチルアミノプロピル)-3-エチルカルボジイミド塩酸塩(648mg、3.38mmol、1.2当量)を添加した。溶液を20℃で0.5時間撹拌し、次いで、N,N-ジメチルホルムアミド(3mL)中の(1S)-1-[4-(1-メチルイミダゾール-2-イル)フェニル]エタンアミン(670mg、2.82mmol、1当量、塩酸塩)およびN,N-ジイソプロピルエチルアミン(1.09g、8.46mmol、1.47mL、3当量)の溶液を、溶液に添加し、11.5時間撹拌した。溶媒を真空中で除去した。粗残渣を半分取逆相HPLC(カラム:Phenomenex Gemini C18 250*50 10um;移動相:[水(0.05%水酸化アンモニアv/v)-ACN];B%:20%~50%、20分;60%分)によって精製した。Tert-ブチル(2S,4R)-4-ヒドロキシ-2-[[(1S)-1-[4-(1-メチルイミダゾール-2-イル)フェニル]エチル]カルバモイル]ピロリジン-1-カルボキシレート(800mg、1.93mmol、収率68%)を無色の油として得た。 To a solution of (2S,4R)-1-tert-butoxycarbonyl-4-hydroxy-pyrrolidine-2-carboxylic acid (651 mg, 2.82 mmol, 1 equiv.) in N,N-dimethylformamide (2 mL) was added 1-hydroxybenzotriazole (456 mg, 3.38 mmol, 1.2 equiv.), 1-(3-dimethylaminopropyl)-3-ethylcarbodiimide hydrochloride (648 mg, 3.38 mmol, 1.2 equiv.). The solution was stirred at 20° C. for 0.5 h, then a solution of (1S)-1-[4-(1-methylimidazol-2-yl)phenyl]ethanamine (670 mg, 2.82 mmol, 1 eq, hydrochloride salt) and N,N-diisopropylethylamine (1.09 g, 8.46 mmol, 1.47 mL, 3 eq) in N,N-dimethylformamide (3 mL) was added to the solution and stirred for 11.5 h. The solvent was removed in vacuo. The crude residue was purified by semi-preparative reverse phase HPLC (column: Phenomenex Gemini C18 250*50 10 um; mobile phase: [water (0.05% ammonia hydroxide v/v)-ACN]; B%: 20%-50%, 20 min; 60% min). Tert-butyl (2S,4R)-4-hydroxy-2-[[(1S)-1-[4-(1-methylimidazol-2-yl)phenyl]ethyl]carbamoyl]pyrrolidine-1-carboxylate (800 mg, 1.93 mmol, 68% yield) was obtained as a colorless oil.
工程5
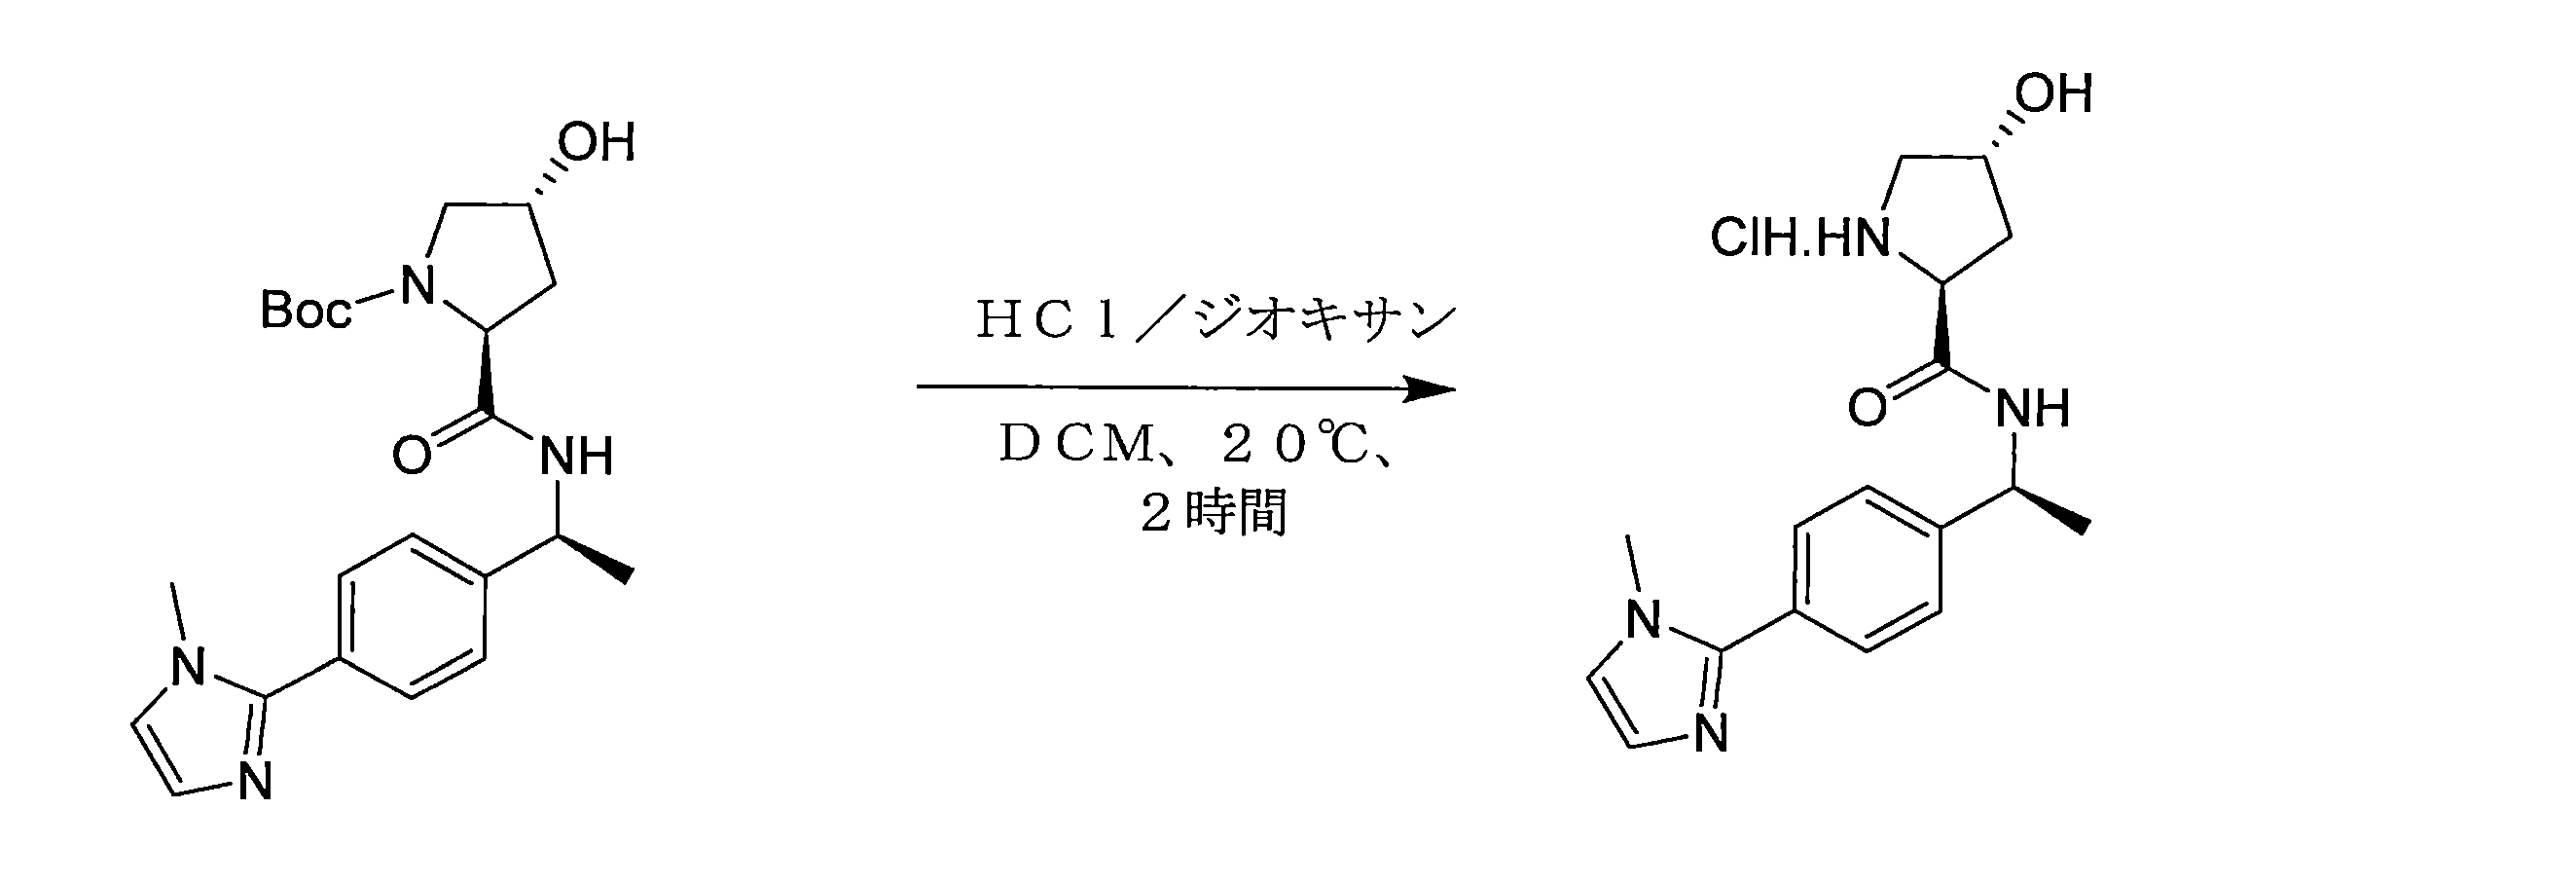
ジクロロメタン(3mL)中のtert-ブチル(2S,4R)-4-ヒドロキシ-2-[[(1S)-1-[4-(1-メチルイミダゾール-2-イル)フェニル]エチル]カルバモイル]ピロリジン-1-カルボキシレート(350mg、0.84mmol、1当量)の溶液に、塩酸/ジオキサン(ジオキサン中4M、3mL、14.21当量)を添加した。次いで、反応混合物を20℃で2時間撹拌した。反応混合物を減圧下で濃縮して、(2S,4R)-4-ヒドロキシ-N-[(1S)-1-[4-(1-メチルイミダゾール-2-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(400mg、粗製、塩酸塩)を淡い黄色の固体として得た。 To a solution of tert-butyl (2S,4R)-4-hydroxy-2-[[(1S)-1-[4-(1-methylimidazol-2-yl)phenyl]ethyl]carbamoyl]pyrrolidine-1-carboxylate (350 mg, 0.84 mmol, 1 equiv.) in dichloromethane (3 mL), hydrochloric acid/dioxane (4 M in dioxane, 3 mL, 14.21 equiv.) was added. The reaction mixture was then stirred at 20°C for 2 h. The reaction mixture was concentrated under reduced pressure to give (2S,4R)-4-hydroxy-N-[(1S)-1-[4-(1-methylimidazol-2-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (400 mg, crude, hydrochloride salt) as a pale yellow solid.
例えば、例示的な化合物47に記載される手順を使用して、(2S,4R)-4-ヒドロキシ-N-[(1S)-1-[4-(1-メチルイミダゾール-2-イル)フェニル]エチル]ピロリジン-2-カルボキサミドを最終化合物に変換した。 For example, (2S,4R)-4-hydroxy-N-[(1S)-1-[4-(1-methylimidazol-2-yl)phenyl]ethyl]pyrrolidine-2-carboxamide was converted to the final compound using the procedure described in exemplary compound 47.
化合物159を、類似の手順を使用して調製した。 Compound 159 was prepared using a similar procedure.
例示的な化合物162の例示的な合成

工程1

250mLの丸底フラスコに、EtOH(60mL)中の1-(6-フルオロピリジン-3-イル)エタン-1-オン(4.17g、29.97mmol、1当量)の溶液を入れた。溶液を水/氷浴中で0℃に冷却し、H2O(10mL)中の水素化ホウ素ナトリウム(2.27g、61.65mmol、2.06当量)の溶液を滴下で添加した。得られた溶液を室温で0.5時間撹拌した。反応混合物を水(400mL)で慎重にクエンチし、酢酸エチル(200mL×3)で抽出した。有機層を合わせ、無水硫酸ナトリウムで乾燥させ、真空下で濃縮した。残渣を、酢酸エチル/石油エーテル(1:1)で溶出するシリカゲルカラム上に適用した。これにより、3.87g(91%)の1-(6-フルオロピリジン-3-イル)エタン-1-オールを淡い黄色の油として得た。 A 250 mL round bottom flask was charged with a solution of 1-(6-fluoropyridin-3-yl)ethan-1-one (4.17 g, 29.97 mmol, 1 equiv.) in EtOH (60 mL). The solution was cooled to 0° C. in a water/ice bath and a solution of sodium borohydride (2.27 g, 61.65 mmol, 2.06 equiv.) in H2O (10 mL) was added dropwise. The resulting solution was stirred at room temperature for 0.5 h. The reaction mixture was carefully quenched with water (400 mL) and extracted with ethyl acetate (200 mL×3). The organic layers were combined, dried over anhydrous sodium sulfate and concentrated under vacuum. The residue was applied onto a silica gel column eluting with ethyl acetate/petroleum ether (1:1). This gave 3.87 g (91%) of 1-(6-fluoropyridin-3-yl)ethan-1-ol as a pale yellow oil.
工程2

窒素不活性雰囲気下でパージし、維持した500mLの三口丸底フラスコに、ジクロロメタン(200mL)中の1-(6-フルオロピリジン-3-イル)エタン-1-オール(4.06g、28.76mmol、1当量)を入れた。溶液を水/氷浴中で0℃に冷却し、PPh3(11.28g、43.01mmol、1.50当量)およびNBS(7.66g、43.04mmol、1.50当量)を0℃で逐次的に添加した。得られた溶液を室温で2時間撹拌した。反応混合液を真空下で濃縮した。残渣を、酢酸エチル/石油エーテル(1:9)で溶出するシリカゲルカラム上に適用した。これにより、5.83g(99%)の5-(1-ブロモエチル)-2-フルオロピリジンを淡い黄色の液体として得た。 A 500 mL three-neck round bottom flask purged and maintained under nitrogen inert atmosphere was charged with 1-(6-fluoropyridin-3-yl)ethan-1-ol (4.06 g, 28.76 mmol, 1 equiv.) in dichloromethane (200 mL). The solution was cooled to 0 °C in a water/ice bath and PPh3 (11.28 g, 43.01 mmol, 1.50 equiv.) and NBS (7.66 g, 43.04 mmol, 1.50 equiv.) were added sequentially at 0 °C. The resulting solution was stirred at room temperature for 2 h. The reaction mixture was concentrated under vacuum. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (1:9). This gave 5.83 g (99%) of 5-(1-bromoethyl)-2-fluoropyridine as a pale yellow liquid.
工程3
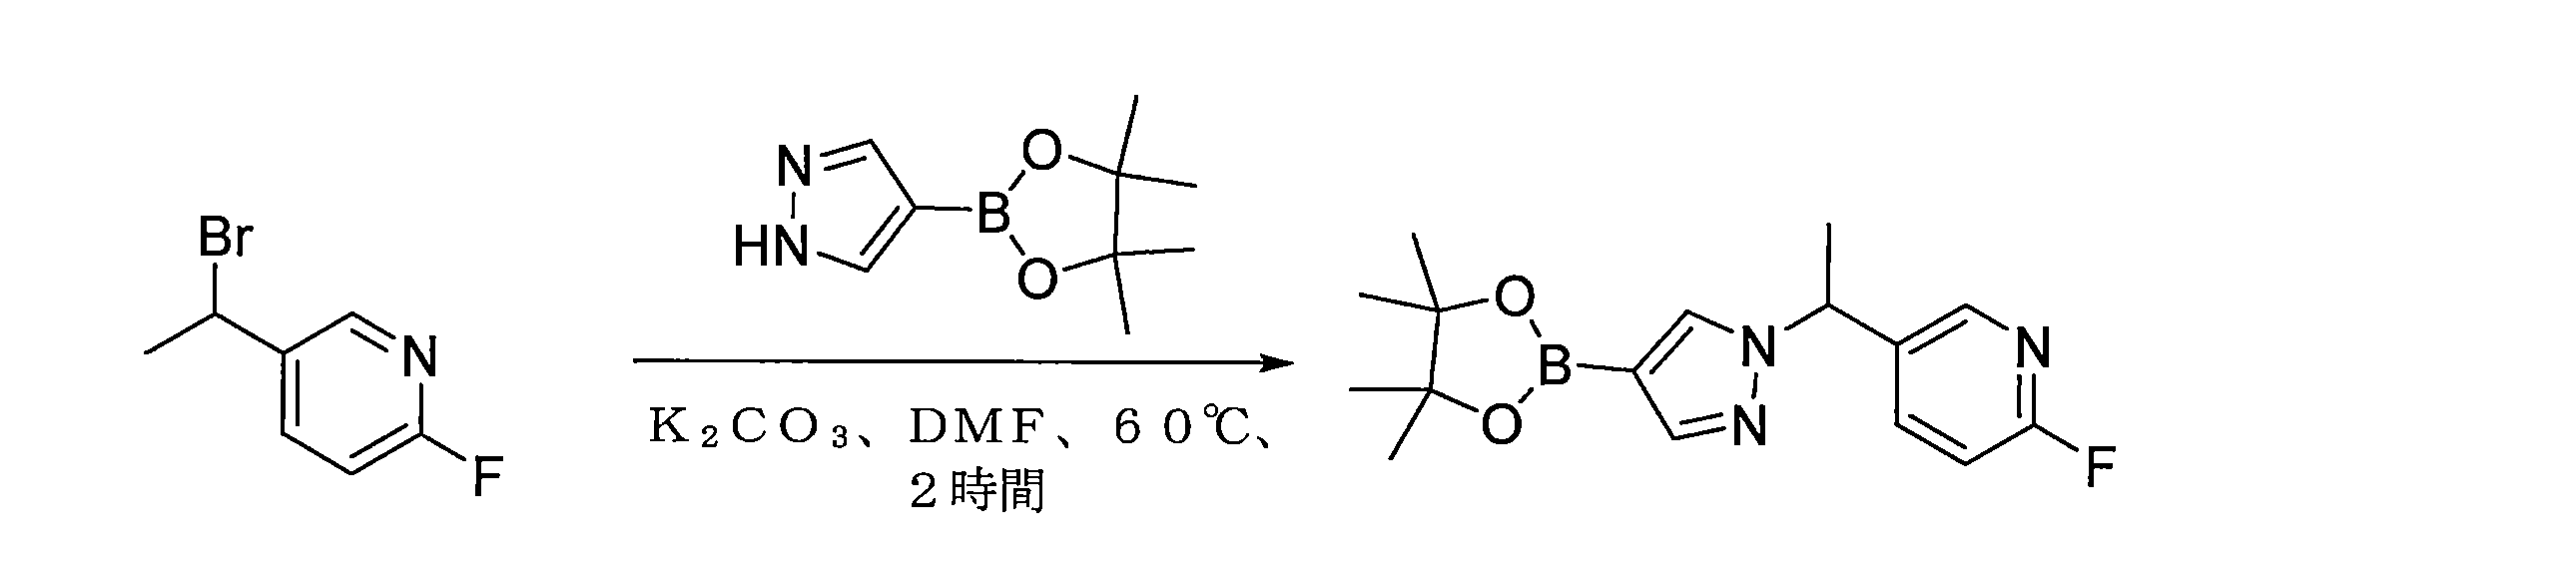
100mLの丸底フラスコに、DMF(50mL)中の5-(1-ブロモエチル)-2-フルオロピリジン(3.06g、15.00mmol、1当量)、4-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)-1H-ピラゾール(3.20g、16.49mmol、1.10当量)、およびK2CO3(4.15g、30.03mmol、2.00当量)の溶液を入れた。得られた混合物を、油浴中で、60℃で2時間撹拌した。反応混合物を室温に冷却し、水(400mL)で希釈した。混合物を酢酸エチル(100mL×4)で抽出し、合わせた有機層をブライン(300mL×2)で洗浄し、無水硫酸ナトリウムで乾燥させ、真空下で濃縮した。これにより、3.43g(72%)の2-フルオロ-5-[1-[4-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)-1H-ピラゾール-1-イル]エチル]ピリジンを淡い黄色の固体として得た。 A 100 mL round-bottom flask was charged with a solution of 5-(1-bromoethyl)-2-fluoropyridine (3.06 g, 15.00 mmol, 1 equiv.), 4-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)-1H-pyrazole (3.20 g, 16.49 mmol, 1.10 equiv.), and K2CO3 (4.15 g, 30.03 mmol, 2.00 equiv.) in DMF (50 mL). The resulting mixture was stirred in an oil bath at 60 °C for 2 h. The reaction mixture was cooled to room temperature and diluted with water (400 mL). The mixture was extracted with ethyl acetate (100 mL × 4) and the combined organic layers were washed with brine (300 mL × 2), dried over anhydrous sodium sulfate, and concentrated under vacuum. This yielded 3.43 g (72%) of 2-fluoro-5-[1-[4-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)-1H-pyrazol-1-yl]ethyl]pyridine as a pale yellow solid.
工程4
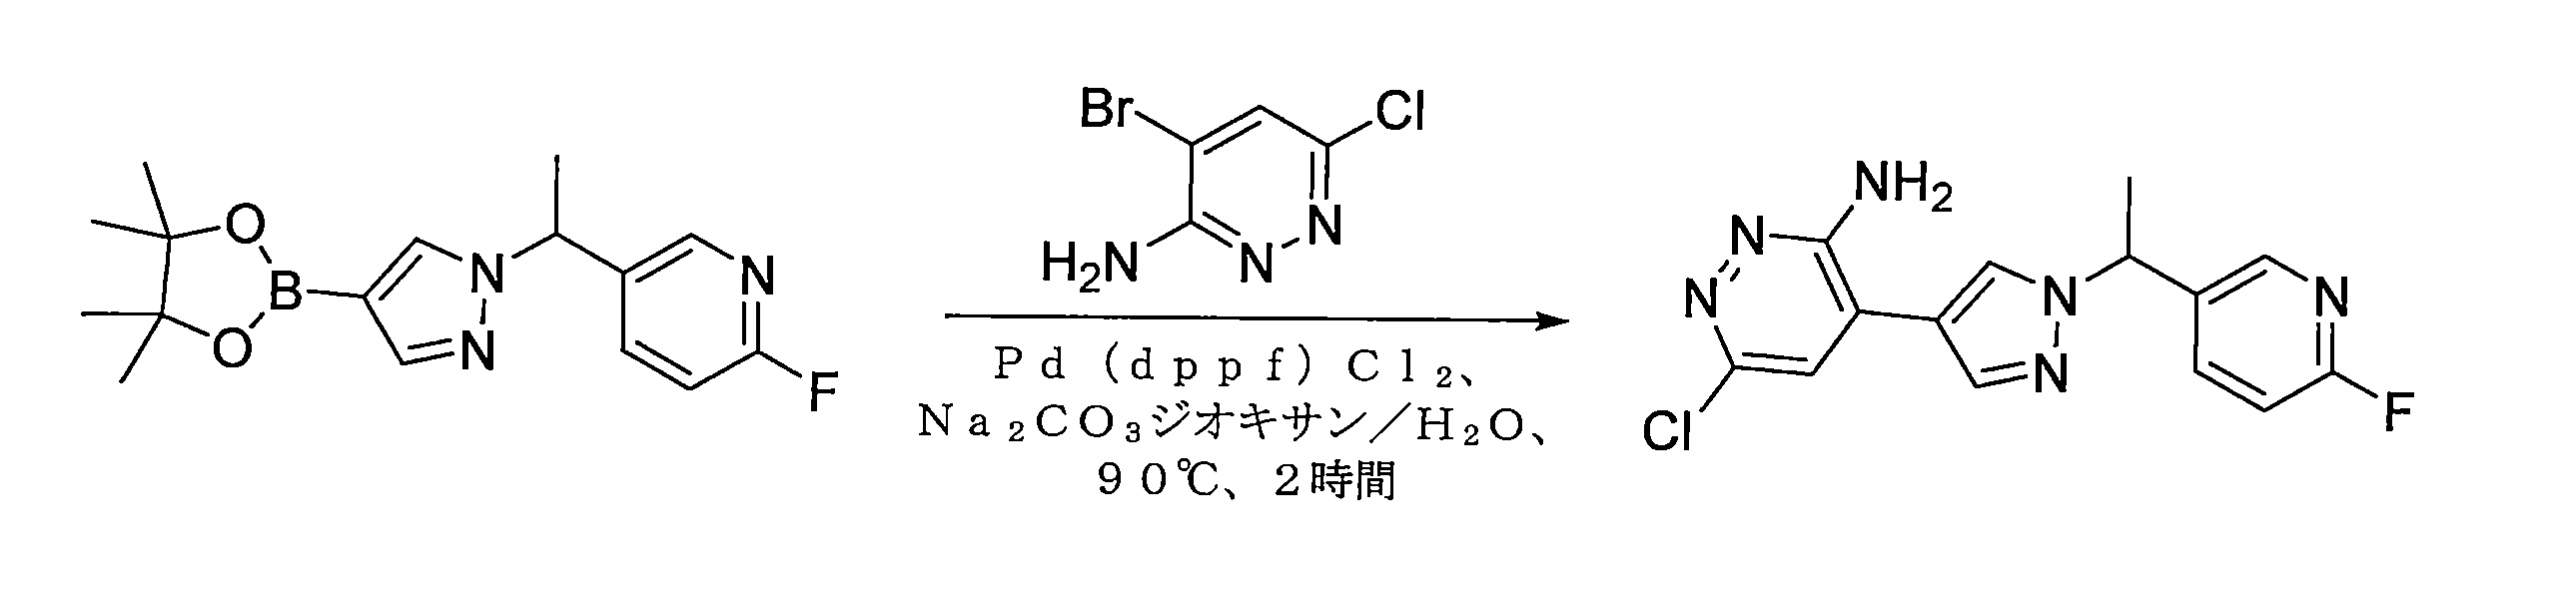
窒素不活性雰囲気下でパージし、維持した100mLの丸底フラスコに、ジオキサン(50mL)およびH2O(8mL)中の2-フルオロ-5-[1-[4-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)-1H-ピラゾール-1-イル]エチル]ピリジン(2.54g、8.01mmol、1当量、4-ブロモ-6-クロロピリダジン-3-アミン(2.00g、9.60mmol、1.20当量)、Na2CO3(2.12g、20.00mmol、2.50当量)、およびPd(dppf)Cl2.CH2Cl2(327mg、0.40mmol、0.05当量)を入れた。得られた混合物を、油浴中で90℃において2時間撹拌した。反応混合物を、室温に冷却し、水(200mL)で希釈した。混合物を酢酸エチル(300mL×2)で抽出し、有機層を合わせ、無水硫酸ナトリウム上で乾燥させ、真空下で濃縮した。残渣を、ジクロロメタン/メタノール(20:1)で溶出するシリカゲルカラム上に適用した。これにより、1.88g(74%)の6-クロロ-4-[1-[1-(6-フルオロピリジン-3-イル)エチル]-1H-ピラゾール-4-イル]ピリダジン-3-アミンを茶色の固体として得た。 In a 100 mL round bottom flask purged and maintained under a nitrogen inert atmosphere, 2-fluoro-5-[1-[4-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)-1H-pyrazol-1-yl]ethyl]pyridine (2.54 g, 8.01 mmol, 1 equiv., 4-bromo-6-chloropyridazin-3-amine (2.00 g, 9.60 mmol, 1.20 equiv.), Na2CO3 (2.12 g, 20.00 mmol, 2.50 equiv.), and Pd(dppf)Cl2.CH2Cl2 (327 mg, 0.40 mmol) in dioxane (50 mL) and H2O (8 mL) were added. mol, 0.05 equiv.) was charged. The resulting mixture was stirred in an oil bath at 90° C. for 2 hours. The reaction mixture was cooled to room temperature and diluted with water (200 mL). The mixture was extracted with ethyl acetate (300 mL×2), and the organic layers were combined, dried over anhydrous sodium sulfate, and concentrated under vacuum. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (20:1). This gave 1.88 g (74%) of 6-chloro-4-[1-[1-(6-fluoropyridin-3-yl)ethyl]-1H-pyrazol-4-yl]pyridazin-3-amine as a brown solid.
工程5
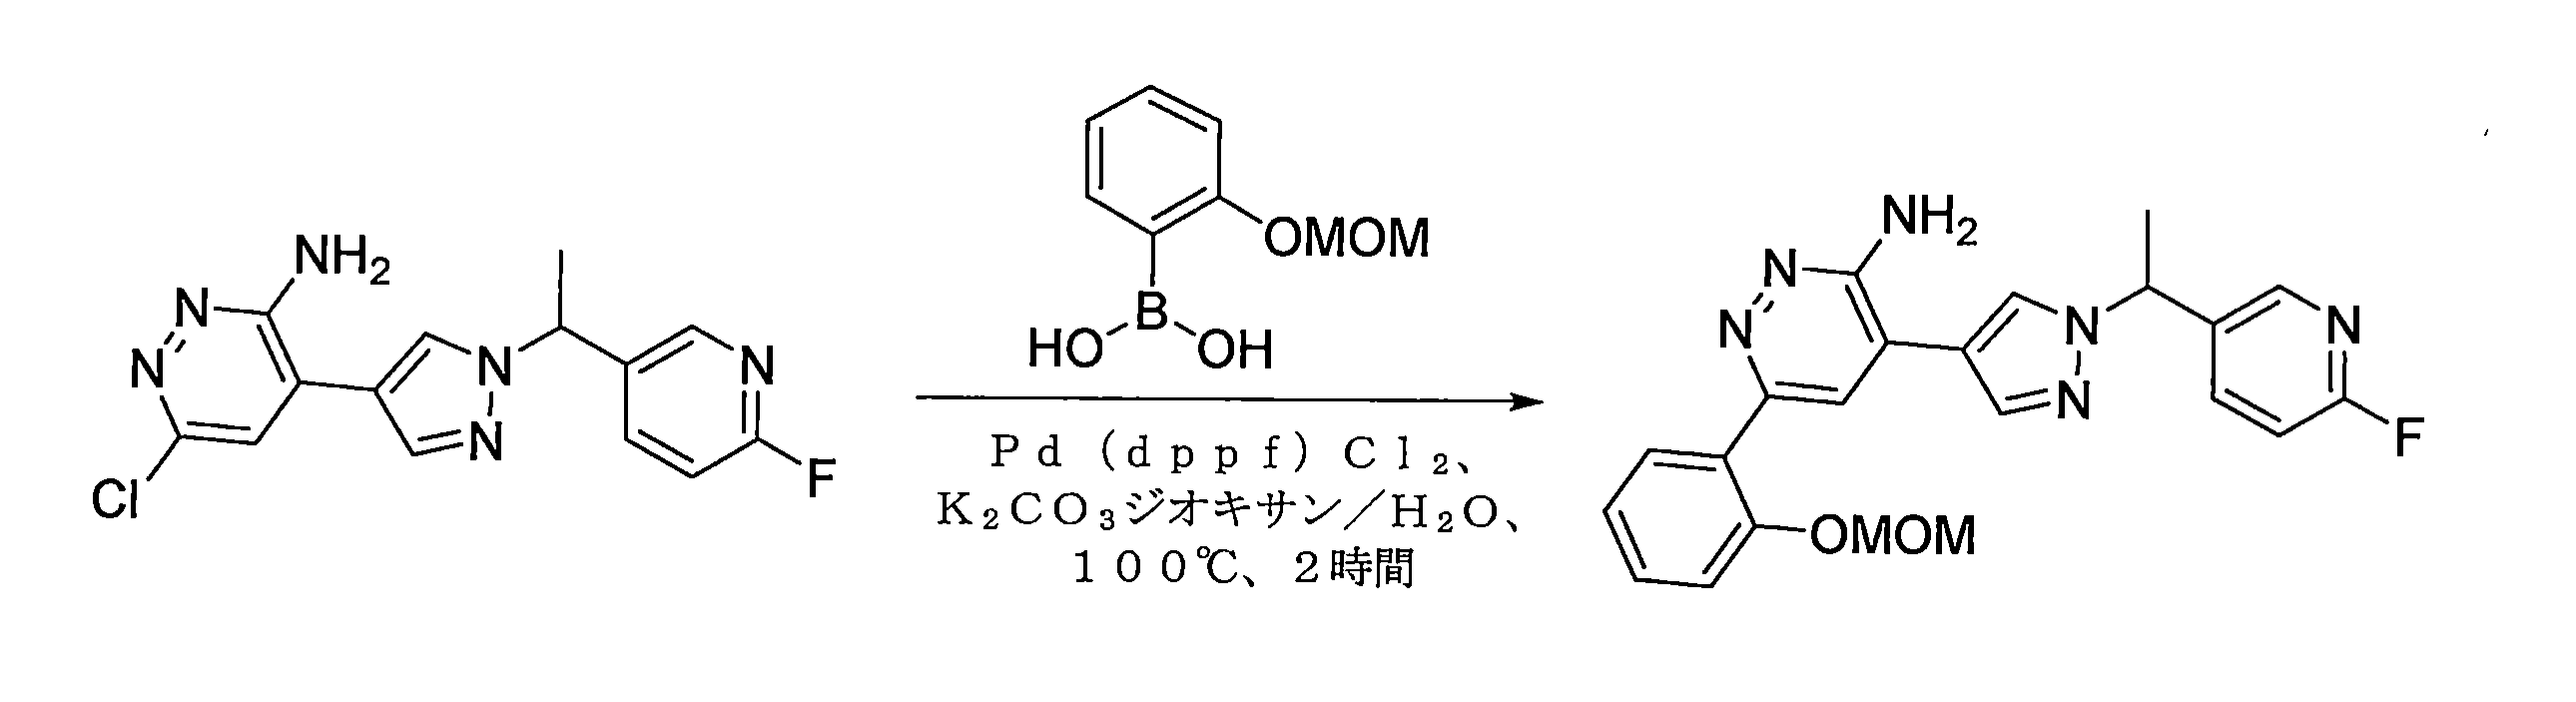
窒素の不活性雰囲気でパージし、維持した30mLの密封チューブに、ジオキサン(18mL)およびH2O(3mL)中の6-クロロ-4-[1-[1-(6-フルオロピリジン-3-イル)エチル]-1H-ピラゾール-4-イル]ピリダジン-3-アミン(765mg、2.40mmol、1当量)、[2-(メトキシメトキシ)フェニル]ボロン酸(524mg、2.88mmol、1.20当量)、K2CO3(829mg、6.00mmol、2.50当量)、およびPd(dppf)Cl2.CH2Cl2(196mg、0.24mmol、0.10当量)を入れた。得られた混合物を、油浴中で、100℃で2時間撹拌した。反応混合物を室温に冷却し、真空下で濃縮した。残渣を、ジクロロメタン/メタノール(20:1)で溶出するシリカゲルカラム上に適用した。これにより、860mg(85%)の4-[1-[1-(6-フルオロピリジン-3-イル)エチル]-1H-ピラゾール-4-イル]-6-[2-(メトキシメトキシ)フェニル]ピリダジン-3-アミンを茶色の固体として得た。 In a 30 mL sealed tube purged and maintained with an inert atmosphere of nitrogen, 6-chloro-4-[1-[1-(6-fluoropyridin-3-yl)ethyl]-1H-pyrazol-4-yl]pyridazin-3-amine (765 mg, 2.40 mmol, 1 equiv.), [2-(methoxymethoxy)phenyl]boronic acid (524 mg, 2.88 mmol, 1.20 equiv.), K2CO3 (829 mg, 6.00 mmol, 2.50 equiv.), and Pd(dppf)Cl2.CH2Cl2 (196 mg, 0.24 mmol, 0.10 equiv.) in dioxane (18 mL) and H2O (3 mL) were placed. The resulting mixture was stirred in an oil bath at 100 °C for 2 h. The reaction mixture was cooled to room temperature and concentrated under vacuum. The residue was applied onto a silica gel column eluting with dichloromethane/methanol (20:1). This yielded 860 mg (85%) of 4-[1-[1-(6-fluoropyridin-3-yl)ethyl]-1H-pyrazol-4-yl]-6-[2-(methoxymethoxy)phenyl]pyridazin-3-amine as a brown solid.
工程6
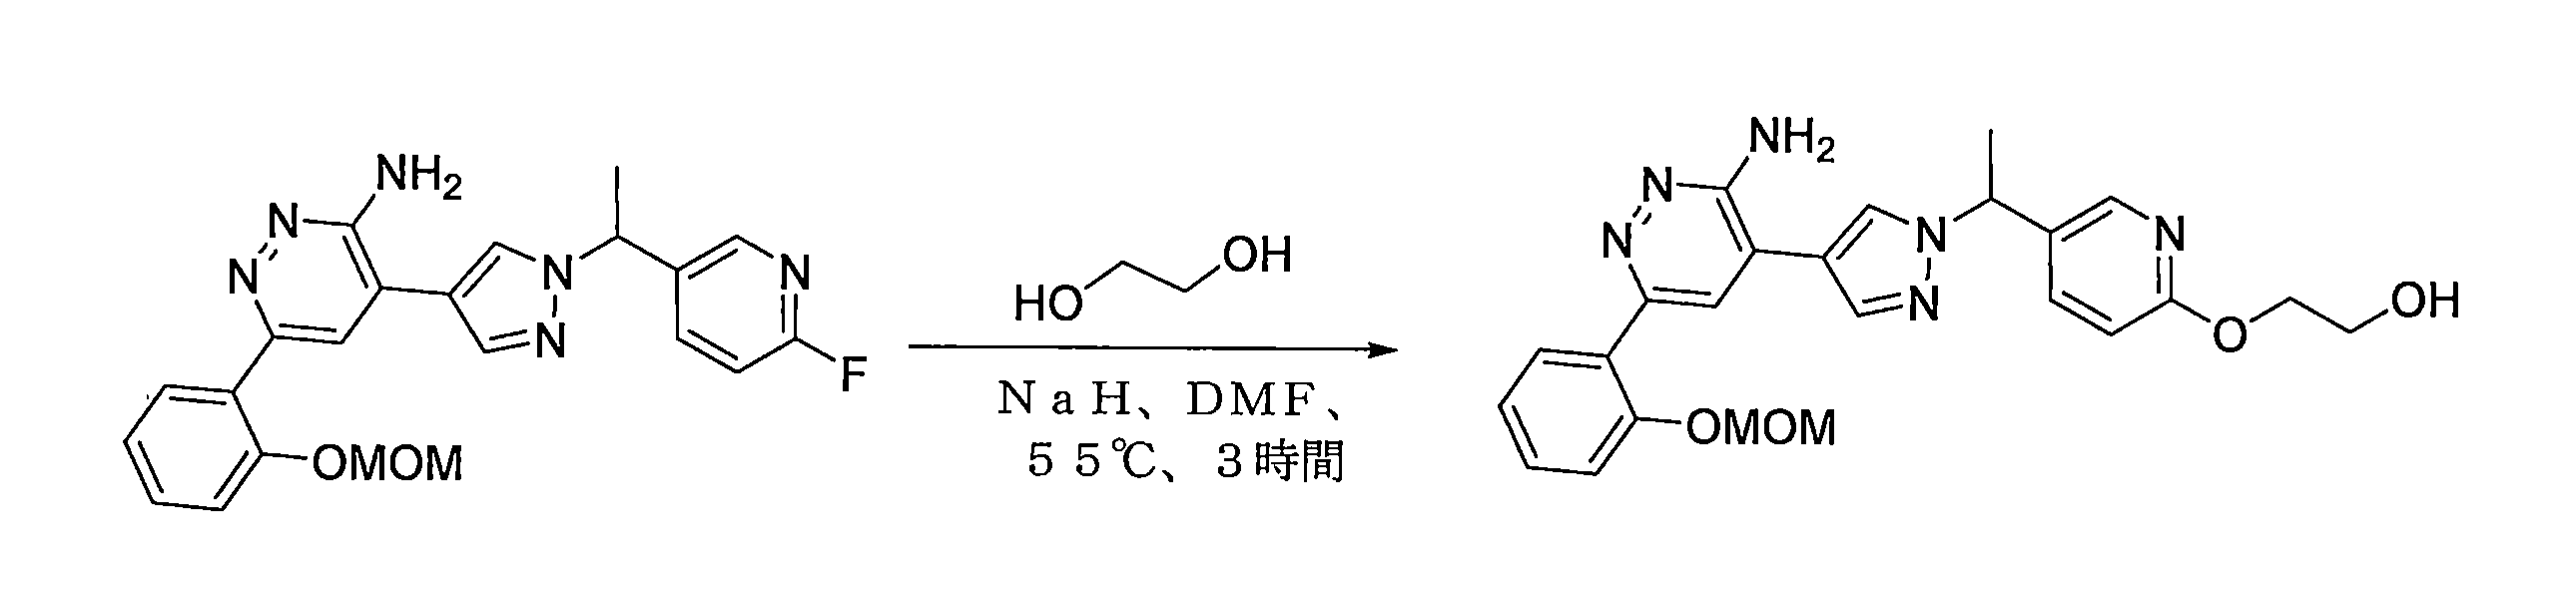
窒素不活性雰囲気下でパージし、維持した50mLの丸底フラスコに、DMF(15mL)中のエタン-1,2-ジオール(1.48g、23.84mmol、10.03当量)を入れた。溶液を水/氷浴中で0℃に冷却し、これにNaH(0.76g、19.00mmol、7.99当量、60%)を0℃で少しずつ添加した。得られた混合物を室温で1.5時間撹拌し、その後、DMF(5mL)中の4-[1-[1-(6-フルオロピリジン-3-イル)エチル]-1H-ピラゾール-4-イル]-6-[2-(メトキシメトキシ)フェニル]ピリダジン-3-アミン(1.00g、2.38mmol、1当量)の溶液を、0℃で、滴下で添加した。温度を油浴中で、55℃で維持しながら、反応混合物をさらに3時間撹拌した。反応混合物を室温に冷却し、次いで、水(200mL)を添加することによってクエンチした。得られた混合物を酢酸エチル(300mL×2)で抽出した。合わせた有機層をブライン(200mL×2)で洗浄し、無水硫酸ナトリウム上で乾燥させ、真空下で濃縮させた。残渣を、ジクロロメタン/メタノール(15:1)で溶出するシリカゲルカラム上に適用した。これにより、361mg(33%)の2-([5-[1-(4-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-1H-ピラゾール-1-イル)エチル]ピリジン-2-イル]オキシ)エタン-1-オールを淡い黄色の固体として得た。 A 50 mL round bottom flask purged and maintained under a nitrogen inert atmosphere was charged with ethane-1,2-diol (1.48 g, 23.84 mmol, 10.03 equiv.) in DMF (15 mL). The solution was cooled to 0°C in a water/ice bath, to which NaH (0.76 g, 19.00 mmol, 7.99 equiv., 60%) was added portionwise at 0°C. The resulting mixture was stirred at room temperature for 1.5 h, after which a solution of 4-[1-[1-(6-fluoropyridin-3-yl)ethyl]-1H-pyrazol-4-yl]-6-[2-(methoxymethoxy)phenyl]pyridazin-3-amine (1.00 g, 2.38 mmol, 1 equiv.) in DMF (5 mL) was added dropwise at 0°C. The reaction mixture was stirred for an additional 3 h while the temperature was maintained at 55°C in an oil bath. The reaction mixture was cooled to room temperature and then quenched by adding water (200 mL). The resulting mixture was extracted with ethyl acetate (300 mL x 2). The combined organic layers were washed with brine (200 mL x 2), dried over anhydrous sodium sulfate, and concentrated under vacuum. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (15:1). This gave 361 mg (33%) of 2-([5-[1-(4-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-1H-pyrazol-1-yl)ethyl]pyridin-2-yl]oxy)ethan-1-ol as a pale yellow solid.
工程7

窒素不活性雰囲気でパージし、維持した50mLの丸底フラスコに、THF(15mL)中のテトラメチルアゾジカルボキサミド(861mg、5.00mmol、10.00当量)およびトリブチルホスフィン(1.01g、4.99mmol、9.98当量)の溶液を0℃で入れた。混合物に、2-([5-[1-(4-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-1H-ピラゾール-1-イル)エチル]ピリジン-2-イル]オキシ)エタン-1-オール(231.3mg、0.50mmol、1当量)および(2S,4R)-4-ヒドロキシ-1-[2-(3-ヒドロキシ-1,2-オキサゾール-5-イル)-3-メチルブタノイル]-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(249mg、0.50mmol、1.00当量)の混合物を添加した。得られた溶液を、0℃で10分間撹拌し、次いで、油浴中で、50℃でさらに2時間撹拌した。反応混合物を室温まで冷却し、水(150mL)で希釈した。得られた混合物を酢酸エチル(150mL)で抽出し、有機層をブライン(150mL×2)で洗浄し、無水硫酸ナトリウムで乾燥させ、真空下で濃縮した。残渣を、ジクロロメタン/メタノール(10/1)で溶出するシリカゲルカラム上に適用した。得られた生成物を、以下の条件で分取HPLCによってさらに精製した。カラム、XBridge Prep OBD C18 Column、19×150mm、5um;移動相、水(10mmol/L NH4HCO3を含む)およびアセトニトリル、35% ACN、8分間で最大53%。これにより、68mg(14%)の(2S,4R)-1-(2-(3-(2-(5-(1-(4-(3-アミノ-6-(2-(メトキシメトキシ)フェニル)ピリダジン-4-イル)-1H-ピラゾール-1-イル)エチル)-2-オキソピリジン-1(2H)-イル)エトキシ)イソキサゾール-5-イル)-3-メチルブタノイル)-4-ヒドロキシ-N-((S)-1-(4-(4-メチルチアゾール-5-イル)フェニル)エチル)ピロリジン-2-カルボキサミドを灰色がかった白色の固体として得た。 A 50 mL round bottom flask purged and maintained with a nitrogen inert atmosphere was charged with a solution of tetramethylazodicarboxamide (861 mg, 5.00 mmol, 10.00 equiv.) and tributylphosphine (1.01 g, 4.99 mmol, 9.98 equiv.) in THF (15 mL) at 0°C. To the mixture was added a mixture of 2-([5-[1-(4-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-1H-pyrazol-1-yl)ethyl]pyridin-2-yl]oxy)ethan-1-ol (231.3 mg, 0.50 mmol, 1 equiv.) and (2S,4R)-4-hydroxy-1-[2-(3-hydroxy-1,2-oxazol-5-yl)-3-methylbutanoyl]-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (249 mg, 0.50 mmol, 1.00 equiv.). The resulting solution was stirred at 0° C. for 10 min and then at 50° C. in an oil bath for an additional 2 h. The reaction mixture was cooled to room temperature and diluted with water (150 mL). The resulting mixture was extracted with ethyl acetate (150 mL), and the organic layer was washed with brine (150 mL x 2), dried over anhydrous sodium sulfate, and concentrated under vacuum. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (10/1). The resulting product was further purified by preparative HPLC with the following conditions: Column, XBridge Prep OBD C18 Column, 19 x 150 mm, 5 um; Mobile phase, water (containing 10 mmol/L NH4HCO3) and acetonitrile, 35% ACN, up to 53% in 8 min. This gave 68 mg (14%) of (2S,4R)-1-(2-(3-(2-(5-(1-(4-(3-amino-6-(2-(methoxymethoxy)phenyl)pyridazin-4-yl)-1H-pyrazol-1-yl)ethyl)-2-oxopyridin-1(2H)-yl)ethoxy)isoxazol-5-yl)-3-methylbutanoyl)-4-hydroxy-N-((S)-1-(4-(4-methylthiazol-5-yl)phenyl)ethyl)pyrrolidine-2-carboxamide as an off-white solid.
上記の他の実施例に記載されるものと類似の手順を使用して、(2S,4R)-1-(2-(3-(2-(5-(1-(4-(3-アミノ-6-(2-(メトキシメトキシ)フェニル)ピリダジン-4-イル)-1H-ピラゾール-1-イル)エチル)-2-オキソピリジン-1(2H)-イル)エトキシ)イソキサゾール-5-イル)-3-メチルブタノイル)-4-ヒドロキシ-N-((S)-1-(4-(4-メチルチアゾール-5-イル)フェニル)エチル)ピロリジン-2-カルボキサミドを最終化合物に変換した。 Using procedures similar to those described in other examples above, (2S,4R)-1-(2-(3-(2-(5-(1-(4-(3-amino-6-(2-(methoxymethoxy)phenyl)pyridazin-4-yl)-1H-pyrazol-1-yl)ethyl)-2-oxopyridin-1(2H)-yl)ethoxy)isoxazol-5-yl)-3-methylbutanoyl)-4-hydroxy-N-((S)-1-(4-(4-methylthiazol-5-yl)phenyl)ethyl)pyrrolidine-2-carboxamide was converted to the final compound.
例示的な化合物157を、類似の手順を使用して調製した。 Exemplary compound 157 was prepared using a similar procedure.
例示的な化合物82の例示的な合成

工程1

100mLの丸底フラスコに、アセトン(20mL)中のメチル2-(3-ヒドロキシ-1,2-オキサゾール-5-イル)-3-メチルブタノエート(498mg、2.50mmol、1当量)、2-(2-ブロモエトキシ)オキサン(627.3mg、3.00mmol、1.20当量)、Cs2CO3(1.63g、5.00mmol、2.00当量)、およびNaI(37.5mg、0.25mmol、0.10当量)を入れた。得られた混合物を、油浴中で、50℃で16時間撹拌した。反応混合物を室温まで冷却し、水(150mL)で希釈した。混合物を酢酸エチル(100mL×2)で抽出し、有機層を合わせ、無水硫酸ナトリウムで乾燥させ、真空下で濃縮した。残渣を、酢酸エチル/石油エーテル(1:9)で溶出するシリカゲルカラム上に適用した。これにより、570mg(70%)のメチル3-メチル-2-[3-[2-(オキサン-2-イルオキシ)エトキシ]-1,2-オキサゾール-5-イル]ブタノエートを淡い黄色の油として得た。 A 100 mL round bottom flask was charged with methyl 2-(3-hydroxy-1,2-oxazol-5-yl)-3-methylbutanoate (498 mg, 2.50 mmol, 1 equiv.), 2-(2-bromoethoxy)oxane (627.3 mg, 3.00 mmol, 1.20 equiv.), Cs2CO3 (1.63 g, 5.00 mmol, 2.00 equiv.), and NaI (37.5 mg, 0.25 mmol, 0.10 equiv.) in acetone (20 mL). The resulting mixture was stirred in an oil bath at 50 °C for 16 h. The reaction mixture was cooled to room temperature and diluted with water (150 mL). The mixture was extracted with ethyl acetate (100 mL × 2), the organic layers were combined, dried over anhydrous sodium sulfate, and concentrated under vacuum. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (1:9). This gave 570 mg (70%) of methyl 3-methyl-2-[3-[2-(oxan-2-yloxy)ethoxy]-1,2-oxazol-5-yl]butanoate as a pale yellow oil.
工程2

100mLの丸底フラスコに、ジオキサン(6mL)中のメチル3-メチル-2-[3-[2-(オキサン-2-イルオキシ)エトキシ]-1,2-オキサゾール-5-イル]ブタノエート(520mg、1.59mmol、1当量)を入れ、それにHCl溶液(ジオキサン中4M、6mL)を室温で添加した。得られた溶液を室温で3時間撹拌した。溶液のpH値を飽和NaHCO3で8に調整した。得られた混合物をジクロロメタン(100mL×4)で抽出し、有機層を合わせ、無水硫酸ナトリウムで乾燥させ、真空下で濃縮した。残渣を、酢酸エチル/石油エーテル(1/1)で溶出するシリカゲルカラム上に適用した。これにより、332mg(86%)のメチル2-[3-(2-ヒドロキシエトキシ)-1,2-オキサゾール-5-イル]-3-メチルブタノエートを淡い黄色の油として得た。 A 100 mL round bottom flask was charged with methyl 3-methyl-2-[3-[2-(oxan-2-yloxy)ethoxy]-1,2-oxazol-5-yl]butanoate (520 mg, 1.59 mmol, 1 equiv.) in dioxane (6 mL), to which HCl solution (4 M in dioxane, 6 mL) was added at room temperature. The resulting solution was stirred at room temperature for 3 h. The pH value of the solution was adjusted to 8 with saturated NaHCO3. The resulting mixture was extracted with dichloromethane (100 mL x 4), the organic layers were combined, dried over anhydrous sodium sulfate and concentrated under vacuum. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (1/1). This gave 332 mg (86%) of methyl 2-[3-(2-hydroxyethoxy)-1,2-oxazol-5-yl]-3-methylbutanoate as a pale yellow oil.
工程3

窒素不活性雰囲気下でパージし、維持した50mLの丸底フラスコに、DMF(6mL)中のメチル2-[3-(2-ヒドロキシエトキシ)-1,2-オキサゾール-5-イル]-3-メチルブタノエート(375mg、1.54mmol、1.20当量)を入れた。溶液を水/氷浴中で0℃に冷却し、NaH(128.4mg、3.21mmol、2.50当量、60%)を添加した。得られた混合物を室温で1時間撹拌し、DMF(4mL)中の4-[1-[1-(6-フルオロピリジン-3-イル)エチル]-1H-ピラゾール-4-イル]-6-[2-(メトキシメトキシ)フェニル]ピリダジン-3-アミン(540mg、1.28mmol、1当量)の溶液を0℃で、滴下で添加した。温度を油浴中で、35℃で維持しながら、得られた混合物をさらに1時間撹拌した。次いで、反応混合物を、飽和NH4Cl溶液(30mL)を添加することによってクエンチし、得られた混合物のpH値を、飽和NaHCO3溶液で約8に調整した。混合物を酢酸エチル(100mL×2)で抽出した。有機層を合わせ、ブライン(100mL×2)で洗浄し、無水硫酸ナトリウムで乾燥させ、真空下で濃縮した。残渣を、ジクロロメタン/メタノール(30/1)で溶出するシリカゲルカラム上に適用した。これにより、540mg(65%)のメチル2-[3-[2-([5-[1-(4-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-1H-ピラゾール-1-イル)エチル]ピリジン-2-イル]オキシ)エトキシ]-1,2-オキサゾール-5-イル]-3-メチルブタノエートを淡い茶色の固体として得た。 A 50 mL round bottom flask purged and maintained under a nitrogen inert atmosphere was charged with methyl 2-[3-(2-hydroxyethoxy)-1,2-oxazol-5-yl]-3-methylbutanoate (375 mg, 1.54 mmol, 1.20 equiv.) in DMF (6 mL). The solution was cooled to 0°C in a water/ice bath and NaH (128.4 mg, 3.21 mmol, 2.50 equiv., 60%) was added. The resulting mixture was stirred at room temperature for 1 h and a solution of 4-[1-[1-(6-fluoropyridin-3-yl)ethyl]-1H-pyrazol-4-yl]-6-[2-(methoxymethoxy)phenyl]pyridazin-3-amine (540 mg, 1.28 mmol, 1 equiv.) in DMF (4 mL) was added dropwise at 0°C. The resulting mixture was stirred for an additional hour while the temperature was maintained at 35°C in an oil bath. The reaction mixture was then quenched by adding saturated NH4Cl solution (30 mL) and the pH value of the resulting mixture was adjusted to about 8 with saturated NaHCO3 solution. The mixture was extracted with ethyl acetate (100 mL x 2). The organic layers were combined, washed with brine (100 mL x 2), dried over anhydrous sodium sulfate and concentrated under vacuum. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (30/1). This gave 540 mg (65%) of methyl 2-[3-[2-([5-[1-(4-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-1H-pyrazol-1-yl)ethyl]pyridin-2-yl]oxy)ethoxy]-1,2-oxazol-5-yl]-3-methylbutanoate as a light brown solid.
上記の他の実施例に記載されるものと類似の手順を使用して、メチル2-[3-[2-([5-[1-(4-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-1H-ピラゾール-1-イル)エチル]ピリジン-2-イル]オキシ)エトキシ]-1,2-オキサゾール-5-イル]-3-メチルブタノエートを最終化合物に変換した。 Using procedures similar to those described in other examples above, methyl 2-[3-[2-([5-[1-(4-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-1H-pyrazol-1-yl)ethyl]pyridin-2-yl]oxy)ethoxy]-1,2-oxazol-5-yl]-3-methylbutanoate was converted to the final compound.
例示的な化合物163を、類似の手順を使用して調製した。 Exemplary compound 163 was prepared using a similar procedure.
例示的な化合物93および例示的な化合物164の例示的な合成

工程1
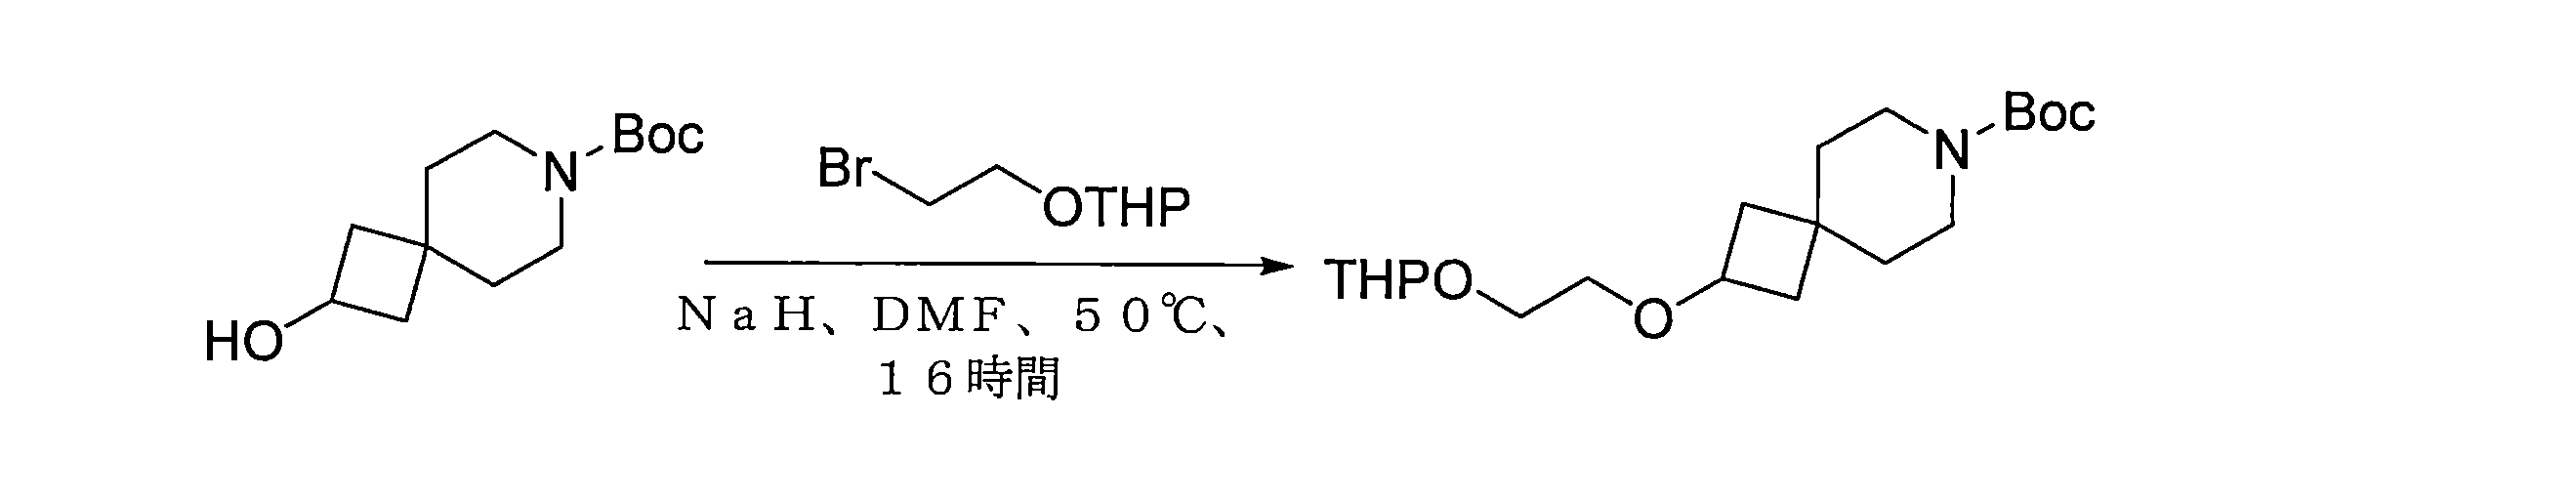
DMF(20mL)中の2-ヒドロキシ-7-アザスピロ[3.5]ノナン-7-カルボキシレート(1.8g、7.46mmol、1当量)の溶液に、水素化ナトリウム(油中60%、358mg、14.9mmol、2当量)を0℃で添加した。混合物を15分間撹拌した。次いで、2-(2-ブロモエトキシ)オキサン(2.3g、11.00mmol、1.47当量)を添加し、混合物を50℃に温め、16時間撹拌した。反応混合物を水(100mL)によってクエンチし、DCM(100mL×3)で抽出し、水(50mL×3)およびブライン(50mL)で洗浄し、無水硫酸ナトリウムで乾燥させ、減圧下で濃縮した。残渣を、酢酸エチル/石油エーテル(1:7)で溶出するシリカゲル上のフラッシュクロマトグラフィーによって精製した。これにより、1.0g(36%)tert-ブチル2-[2-(オキサン-2-イルオキシ)エトキシ]-7-アザスピロ[3.5]ノナン-7-カルボキシレートを無色の油として得た。 To a solution of 2-hydroxy-7-azaspiro[3.5]nonane-7-carboxylate (1.8 g, 7.46 mmol, 1 eq.) in DMF (20 mL) was added sodium hydride (60% in oil, 358 mg, 14.9 mmol, 2 eq.) at 0 °C. The mixture was stirred for 15 min. Then 2-(2-bromoethoxy)oxane (2.3 g, 11.00 mmol, 1.47 eq.) was added and the mixture was warmed to 50 °C and stirred for 16 h. The reaction mixture was quenched by water (100 mL), extracted with DCM (100 mL x 3), washed with water (50 mL x 3) and brine (50 mL), dried over anhydrous sodium sulfate and concentrated under reduced pressure. The residue was purified by flash chromatography on silica gel eluting with ethyl acetate/petroleum ether (1:7). This yielded 1.0 g (36%) tert-butyl 2-[2-(oxan-2-yloxy)ethoxy]-7-azaspiro[3.5]nonane-7-carboxylate as a colorless oil.
工程2

100mLの丸底フラスコに、ジオキサン(15mL)中のtert-ブチル2-[2-(オキサン-2-イルオキシ)エトキシ]-7-アザスピロ[3.5]ノナン-7-カルボキシレート(1.0g、2.71mmol、1当量)の溶液を入れ、それに1,4-ジオキサン溶液(4.0M、15mL)中の塩化水素を添加した。得られた溶液を室温で16時間撹拌し、次いで、減圧下で濃縮した。これにより、800mg(53%)の2-[7-アザスピロ[3.5]ノナン-2-イルオキシ]エタン-1-オールを黄色の油として得た。 A 100 mL round bottom flask was charged with a solution of tert-butyl 2-[2-(oxan-2-yloxy)ethoxy]-7-azaspiro[3.5]nonane-7-carboxylate (1.0 g, 2.71 mmol, 1 equiv.) in dioxane (15 mL) to which was added hydrogen chloride in 1,4-dioxane solution (4.0 M, 15 mL). The resulting solution was stirred at room temperature for 16 hours and then concentrated under reduced pressure. This gave 800 mg (53%) of 2-[7-azaspiro[3.5]nonan-2-yloxy]ethan-1-ol as a yellow oil.
工程3

100mLの丸底フラスコに、DCM(30.0mL)中の2-[7-アザスピロ[3.5]ノナン-2-イルオキシ]エタン-1-オール(600mg、3.24mmol、1当量)、Et3N(983mg、9.72mmol、3当量)を入れ、それにCbzCl(663mg、3.89mmol、1.2当量)を室温で添加した。得られた溶液を室温で16時間撹拌した。次いで、反応混合物を50mLの水で希釈し、酢酸エチル(50mL×2)で抽出した。有機相を合わせ、ブラインで洗浄し、硫酸ナトリウムで乾燥させ、減圧下で濃縮した。残渣を、ジクロロメタン/メタノール(v:v=10:1)で溶出するシリカゲルカラム上に適用した。これにより、210mg(20%)のベンジル2-(2-ヒドロキシエトキシ)-7-アザスピロ[3.5]ノナン-7-カルボキシレートを黄色の油として得た。 A 100 mL round bottom flask was charged with 2-[7-azaspiro[3.5]nonan-2-yloxy]ethan-1-ol (600 mg, 3.24 mmol, 1 eq.), Et3N (983 mg, 9.72 mmol, 3 eq.) in DCM (30.0 mL), to which CbzCl (663 mg, 3.89 mmol, 1.2 eq.) was added at room temperature. The resulting solution was stirred at room temperature for 16 hours. The reaction mixture was then diluted with 50 mL of water and extracted with ethyl acetate (50 mL x 2). The organic phases were combined, washed with brine, dried over sodium sulfate, and concentrated under reduced pressure. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (v:v=10:1). This afforded 210 mg (20%) of benzyl 2-(2-hydroxyethoxy)-7-azaspiro[3.5]nonane-7-carboxylate as a yellow oil.
工程4

窒素不活性雰囲気下でパージし、維持した50mLの丸底フラスコに、DCM(20mL)中のベンジル2-(2-ヒドロキシエトキシ)-7-アザスピロ[3.5]ノナン-7-カルボキシレート(400mg、1.25mmol、1当量)、Et3N(380mg、3.76mmol、3当量)を入れ、それにMsCl(287mg、2.50mmol、2.00当量)を0℃で添加した。得られた混合物を水/氷浴中で、0℃において1時間撹拌した。反応混合物を、水(30mL)を添加することによってクエンチし、ジクロロメタン(30mL×2)で抽出した。合わせた有機層をブライン(30mL)で洗浄し、無水硫酸ナトリウムで乾燥させ、濃縮した。これにより、420mg(85%)のベンジル2-[2-(メタンスルホニルオキシ)エトキシ]-7-アザスピロ[3.5]ノナン-7-カルボキシレートを黄色の油として得た。 A 50 mL round bottom flask purged and maintained under nitrogen inert atmosphere was charged with benzyl 2-(2-hydroxyethoxy)-7-azaspiro[3.5]nonane-7-carboxylate (400 mg, 1.25 mmol, 1 eq.), Et3N (380 mg, 3.76 mmol, 3 eq.) in DCM (20 mL) to which MsCl (287 mg, 2.50 mmol, 2.00 eq.) was added at 0°C. The resulting mixture was stirred in a water/ice bath at 0°C for 1 h. The reaction mixture was quenched by adding water (30 mL) and extracted with dichloromethane (30 mL x 2). The combined organic layers were washed with brine (30 mL), dried over anhydrous sodium sulfate, and concentrated. This gave 420 mg (85%) of benzyl 2-[2-(methanesulfonyloxy)ethoxy]-7-azaspiro[3.5]nonane-7-carboxylate as a yellow oil.
工程5

100mLの丸底フラスコに、MeCN(15mL)中のベンジル2-[2-(メタンスルホニルオキシ)エトキシ]-7-アザスピロ[3.5]ノナン-7-カルボキシレート(300mg、0.75mmol、1当量)、6-[2-(メトキシメトキシ)フェニル]-4-[1-オキサ-4,9-ジアザスピロ[5.5]ウンデカン-9-イル]ピリダジン-3-アミン(291mg、0.75mmol、1当量)、K2CO3(313mg、2.26mmol、3当量)、NaI(113mg、0.75mmol、1当量)の溶液を入れた。得られた混合物を、油浴中で、75℃で16時間撹拌した。固形分を濾過し、濾液を減圧下で濃縮した。残渣を、ジクロロメタン/メタノール(v:v=10:1)で溶出するシリカゲルカラム上に適用した。これにより、130mg(25%)のベンジル2-[2-(9-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-1-オキサ-4,9-ジアザスピロ[5.5]ウンデカン-4-イル)エトキシ]-7-アザスピロ[3.5]ノナン-7-カルボキシレートを黄色の油として得た。 A 100 mL round bottom flask was charged with a solution of benzyl 2-[2-(methanesulfonyloxy)ethoxy]-7-azaspiro[3.5]nonane-7-carboxylate (300 mg, 0.75 mmol, 1 equiv.), 6-[2-(methoxymethoxy)phenyl]-4-[1-oxa-4,9-diazaspiro[5.5]undecan-9-yl]pyridazin-3-amine (291 mg, 0.75 mmol, 1 equiv.), K2CO3 (313 mg, 2.26 mmol, 3 equiv.), NaI (113 mg, 0.75 mmol, 1 equiv.) in MeCN (15 mL). The resulting mixture was stirred in an oil bath at 75 °C for 16 h. The solids were filtered and the filtrate was concentrated under reduced pressure. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (v:v=10:1). This gave 130 mg (25%) of benzyl 2-[2-(9-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-1-oxa-4,9-diazaspiro[5.5]undecan-4-yl)ethoxy]-7-azaspiro[3.5]nonane-7-carboxylate as a yellow oil.
工程6
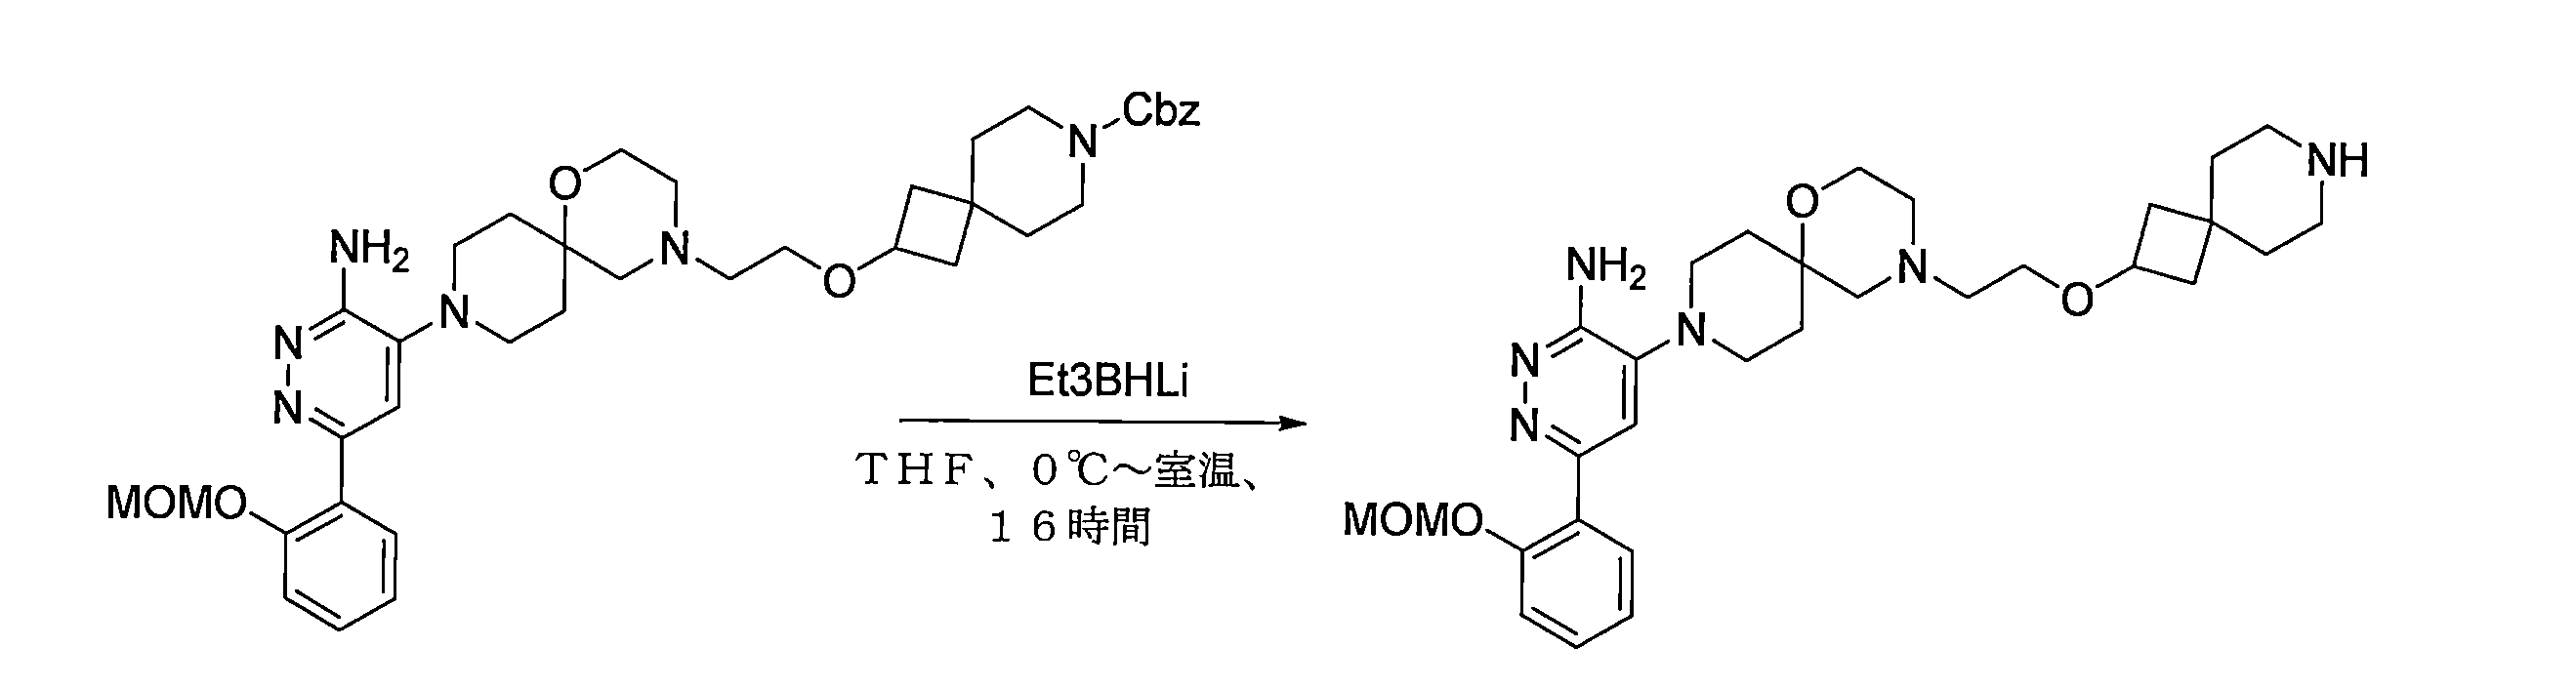
100mLの丸底フラスコに、THF(5mL)中のベンジル2-[2-(9-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-1-オキサ-4,9-ジアザスピロ[5.5]ウンデカン-4-イル)エトキシ]-7-アザスピロ[3.5]ノナン-7-カルボキシレート(230mg、0.33mmol、1当量)の溶液を入れ、それに水素化トリエチルホウ素リチウム(THF中1.0M、1.65mmol、5.0当量)を0℃で添加した。得られた混合物を室温で16時間撹拌し、次いで、水(50mL)によってクエンチし、ジクロロメタン(50mL×3)で抽出した。合わせた有機層をブライン(50mL)で洗浄し、無水硫酸ナトリウムで乾燥させ、減圧下で濃縮した。これにより、140mg(76%)の4-[4-(2-[7-アザスピロ[3.5]ノナン-2-イルオキシ]エチル)-1-オキサ-4,9-ジアザスピロ[5.5]ウンデカン-9-イル]-6-[2-(メトキシメトキシ)フェニル]ピリダジン-3-アミンを黄色の油として得た。 A 100 mL round bottom flask was charged with a solution of benzyl 2-[2-(9-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-1-oxa-4,9-diazaspiro[5.5]undecan-4-yl)ethoxy]-7-azaspiro[3.5]nonane-7-carboxylate (230 mg, 0.33 mmol, 1 equiv.) in THF (5 mL), to which lithium triethylborohydride (1.0 M in THF, 1.65 mmol, 5.0 equiv.) was added at 0°C. The resulting mixture was stirred at room temperature for 16 h, then quenched by water (50 mL) and extracted with dichloromethane (50 mL x 3). The combined organic layers were washed with brine (50 mL), dried over anhydrous sodium sulfate, and concentrated under reduced pressure. This gave 140 mg (76%) of 4-[4-(2-[7-azaspiro[3.5]nonan-2-yloxy]ethyl)-1-oxa-4,9-diazaspiro[5.5]undecan-9-yl]-6-[2-(methoxymethoxy)phenyl]pyridazin-3-amine as a yellow oil.
上記の他の実施例について記載される手順を使用して、以下のスキームに従って、4-[4-(2-[7-アザスピロ[3.5]ノナン-2-イルオキシ]エチル)-1-オキサ-4,9-ジアザスピロ[5.5]ウンデカン-9-イル]-6-[2-(メトキシメトキシ)フェニル]ピリダジン-3-アミンを最終化合物に変換した。

例示的な化合物164を、類似の手順を使用して調製した。 Exemplary compound 164 was prepared using a similar procedure.
例示的な化合物113の例示的な合成

工程1
ジクロロメタン(200mL)中のブタ-2-イン-1,4-ジオール(10g、116.16mmol、1当量)、2,3-ジヒドロ-2H-ピラン(9.77g、116.16mmol、10.6mL、1当量)の溶液に、p-トルエンスルホン酸ピリジニウム(2.92g、11.62mmol、0.1当量)を添加した。反応混合物を40℃で1時間撹拌した。水(200mL)を混合物に添加し、得られた混合物をジクロロメタン(150mL×2)で抽出した。合わせた有機相をブライン(150mL)で洗浄し、硫酸ナトリウムで乾燥させ、濾過し、真空中で濃縮した。残渣を、シリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=100:1~1:1)によって精製した。4-テトラヒドロピラン-2-イルオキシブタ-2-イン-1-オール(10.2g、59.93mmol、収率51%)を淡い黄色の油として得た。 To a solution of but-2-yn-1,4-diol (10 g, 116.16 mmol, 1 eq.), 2,3-dihydro-2H-pyran (9.77 g, 116.16 mmol, 10.6 mL, 1 eq.) in dichloromethane (200 mL), pyridinium p-toluenesulfonate (2.92 g, 11.62 mmol, 0.1 eq.) was added. The reaction mixture was stirred at 40 °C for 1 h. Water (200 mL) was added to the mixture, and the resulting mixture was extracted with dichloromethane (150 mL x 2). The combined organic phase was washed with brine (150 mL), dried over sodium sulfate, filtered, and concentrated in vacuum. The residue was purified by silica gel chromatography (petroleum ether: ethyl acetate = 100: 1 to 1: 1). 4-Tetrahydropyran-2-yloxybut-2-yn-1-ol (10.2 g, 59.93 mmol, 51% yield) was obtained as a pale yellow oil.
工程2

ジクロロメタン(100mL)中の4-テトラヒドロピラン-2-イルオキシブタ-2-イン-1-オール(5g、29.38mmol、1当量)、トリフェニルホスフィン(7.70g、29.38mmol、1当量)の溶液に、四臭化炭素(10.72g、32.31mmol、1.1当量)を0℃で添加した。反応混合物を20℃で1時間撹拌した。混合物を真空中で濃縮した。残渣を、シリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=1:0~100:1)によって精製した。2-(4-ブロモブタ-2-イノキシ)テトラヒドロピラン(4g、17.16mmol、収率58%)を淡い黄色の油として得た。 To a solution of 4-tetrahydropyran-2-yloxybut-2-yn-1-ol (5 g, 29.38 mmol, 1 eq.), triphenylphosphine (7.70 g, 29.38 mmol, 1 eq.) in dichloromethane (100 mL), carbon tetrabromide (10.72 g, 32.31 mmol, 1.1 eq.) was added at 0 °C. The reaction mixture was stirred at 20 °C for 1 h. The mixture was concentrated in vacuum. The residue was purified by silica gel chromatography (petroleum ether: ethyl acetate = 1:0 to 100:1). 2-(4-bromobut-2-ynoxy)tetrahydropyran (4 g, 17.16 mmol, 58% yield) was obtained as a pale yellow oil.
工程3

アセトニトリル(30mL)中の4-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)-1H-ピラゾール(3g、15.46mmol、1当量)、2-(4-ブロモブタ-2-イノキシ)テトラヒドロピラン(3.96g、17.01mmol、1.1当量)の溶液に、炭酸カリウム(2.56g、18.55mmol、1.2当量)を添加した。反応混合物を70℃で12時間撹拌した。水(100mL)を混合物に添加し、得られた混合物を酢酸エチル(30mL×3)で抽出した。合わせた有機相をブライン(50mL)で洗浄し、硫酸ナトリウムで乾燥させ、濾過し、真空中で濃縮した。残渣を、シリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=200:1~3:1)によって精製した。1-(4-テトラヒドロピラン-2-イルオキシブタ-2-イニル)-4-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)ピラゾール(4.5g、13.00mmol、収率84%)を淡い黄色の油として得た。 To a solution of 4-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)-1H-pyrazole (3 g, 15.46 mmol, 1 eq.), 2-(4-bromobut-2-ynoxy)tetrahydropyran (3.96 g, 17.01 mmol, 1.1 eq.) in acetonitrile (30 mL), potassium carbonate (2.56 g, 18.55 mmol, 1.2 eq.) was added. The reaction mixture was stirred at 70 °C for 12 h. Water (100 mL) was added to the mixture, and the resulting mixture was extracted with ethyl acetate (30 mL x 3). The combined organic phase was washed with brine (50 mL), dried over sodium sulfate, filtered, and concentrated in vacuum. The residue was purified by silica gel chromatography (petroleum ether: ethyl acetate = 200: 1 to 3: 1). 1-(4-tetrahydropyran-2-yloxybut-2-ynyl)-4-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)pyrazole (4.5 g, 13.00 mmol, 84% yield) was obtained as a pale yellow oil.
工程4

ジオキサン(60mL)および水(20mL)中の1-(4-テトラヒドロピラン-2-イルオキシブタ-2-イニル)-4-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)ピラゾール(2g、5.78mmol、1当量)、4-ブロモ-6-クロロ-ピリダジン-3-アミン(1.20g、5.78mmol、1当量)の溶液に、炭酸ナトリウム(1.53g、14.44mmol、2.5当量)および(1,1’-ビス(ジフェニルホスフィノ)フェロセン)パラジウム(II)二塩化物(422mg、0.58mmol、0.1当量)を窒素下で添加した。反応混合物を110℃で3時間撹拌した。水(200mL)を混合物に添加し、得られた混合物を酢酸エチル(100mL×3)で抽出した。合わせた有機相をブライン(100mL)で洗浄し、硫酸ナトリウムで乾燥させ、濾過し、真空中で濃縮した。残渣を、シリカゲルクロマトグラフィー(ジクロロメタン:メタノール=1:0~50:1)によって精製した。6-クロロ-4-[1-(4-テトラヒドロピラン-2-イルオキシブタ-2-イニル)ピラゾール-4-イル]ピリダジン-3-アミン(1.3g、3.40mmol、収率58%)を黄色の固体として得た。 To a solution of 1-(4-tetrahydropyran-2-yloxybut-2-ynyl)-4-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)pyrazole (2 g, 5.78 mmol, 1 eq.), 4-bromo-6-chloro-pyridazin-3-amine (1.20 g, 5.78 mmol, 1 eq.) in dioxane (60 mL) and water (20 mL), sodium carbonate (1.53 g, 14.44 mmol, 2.5 eq.) and (1,1'-bis(diphenylphosphino)ferrocene)palladium(II) dichloride (422 mg, 0.58 mmol, 0.1 eq.) were added under nitrogen. The reaction mixture was stirred at 110 °C for 3 h. Water (200 mL) was added to the mixture and the resulting mixture was extracted with ethyl acetate (100 mL x 3). The combined organic phase was washed with brine (100 mL), dried over sodium sulfate, filtered, and concentrated in vacuo. The residue was purified by silica gel chromatography (dichloromethane:methanol=1:0 to 50:1). 6-chloro-4-[1-(4-tetrahydropyran-2-yloxybut-2-ynyl)pyrazol-4-yl]pyridazin-3-amine (1.3 g, 3.40 mmol, 58% yield) was obtained as a yellow solid.
工程5

メタノール(15mL)中の6-クロロ-4-[1-(4-テトラヒドロピラン-2-イルオキシブタ-2-イニル)ピラゾール-4-イル]ピリダジン-3-アミン(1.6g、4.60mmol、1当量)の溶液に、塩酸溶液(メタノール中4M、15mL、13.04当量)を添加した。反応混合物を20℃で1時間撹拌した。飽和重炭酸ナトリウム(50mL)を混合物に添加してpHを約6に調整し、その後、得られた混合物を酢酸エチル(30mL×3)で抽出した。合わせた有機相を、ブライン(30mL)で洗浄し、硫酸ナトリウム上で乾燥させ、濾過し、真空中で濃縮した。4-[4-(3-アミノ-6-クロロ-ピリダジン-4-イル)ピラゾール-1-イル]ブタ-2-イン-1-オール(1.0g、3.79mmol、収率82%)を淡い黄色の固体として得た。 To a solution of 6-chloro-4-[1-(4-tetrahydropyran-2-yloxybut-2-ynyl)pyrazol-4-yl]pyridazin-3-amine (1.6 g, 4.60 mmol, 1 eq.) in methanol (15 mL) was added hydrochloric acid solution (4 M in methanol, 15 mL, 13.04 eq.). The reaction mixture was stirred at 20°C for 1 h. Saturated sodium bicarbonate (50 mL) was added to the mixture to adjust the pH to about 6, and then the resulting mixture was extracted with ethyl acetate (30 mL x 3). The combined organic phase was washed with brine (30 mL), dried over sodium sulfate, filtered, and concentrated in vacuo. 4-[4-(3-amino-6-chloro-pyridazin-4-yl)pyrazol-1-yl]but-2-yn-1-ol (1.0 g, 3.79 mmol, 82% yield) was obtained as a pale yellow solid.
工程6

テトラヒドロフラン(20mL)中の4-[4-(3-アミノ-6-クロロ-ピリダジン-4-イル)ピラゾール-1-イル]ブタ-2-イン-1-オール(1.0g、3.79mmol、1当量)、トリフェニルホスフィン(1.19g、4.55mmol、1.2当量)の溶液に、四臭化炭素(1.51g、4.55mmol、1.2当量)を添加した。反応混合物を20℃で1時間撹拌した。混合物を真空中で濃縮した。残渣を、シリカゲルクロマトグラフィー(ジクロロメタン:メタノール=1:0~50:1)によって精製した。4-[1-(4-ブロモブタ-2-イニル)ピラゾール-4-イル]-6-クロロ-ピリダジン-3-アミン(1.0g、1.87mmol、収率49%)を淡い黄色の固体として得た。 To a solution of 4-[4-(3-amino-6-chloro-pyridazin-4-yl)pyrazol-1-yl]but-2-yn-1-ol (1.0 g, 3.79 mmol, 1 eq.), triphenylphosphine (1.19 g, 4.55 mmol, 1.2 eq.) in tetrahydrofuran (20 mL), carbon tetrabromide (1.51 g, 4.55 mmol, 1.2 eq.) was added. The reaction mixture was stirred at 20°C for 1 h. The mixture was concentrated in vacuum. The residue was purified by silica gel chromatography (dichloromethane:methanol = 1:0 to 50:1). 4-[1-(4-bromobut-2-ynyl)pyrazol-4-yl]-6-chloro-pyridazin-3-amine (1.0 g, 1.87 mmol, 49% yield) was obtained as a pale yellow solid.
工程7

アセトニトリル(10mL)中の4-[1-(4-ブロモブタ-2-イニル)ピラゾール-4-イル]-6-クロロ-ピリダジン-3-アミン(1g、1.87mmol、1当量)およびtert-ブチルピペラジン-1-カルボキシレート(696mg、3.74mmol、2当量)の溶液に、N,N-ジイソプロピルエチルアミン(724mg、5.60mmol、1.0mL、3当量)を添加した。混合物を80℃で12時間撹拌した。水(30mL)を混合物に添加し、得られた混合物を酢酸エチル(20mL×3)で抽出した。合わせた有機相をブライン(20mL)で洗浄し、硫酸ナトリウムで乾燥させ、濾過し、真空中で濃縮した。残渣を、シリカゲルクロマトグラフィー(ジクロロメタン:メタノール=1:0~20:1)によって精製した。Tert-ブチル4-[4-[4-(3-アミノ-6-クロロ-ピリダジン-4-イル)ピラゾール-1-イル]ブタ-2-イニル]ピペラジン-1-カルボキシレート(600mg、1.34mmol、収率71%)を茶色の固体として得た。 To a solution of 4-[1-(4-bromobut-2-ynyl)pyrazol-4-yl]-6-chloro-pyridazin-3-amine (1 g, 1.87 mmol, 1 eq.) and tert-butyl piperazine-1-carboxylate (696 mg, 3.74 mmol, 2 eq.) in acetonitrile (10 mL), N,N-diisopropylethylamine (724 mg, 5.60 mmol, 1.0 mL, 3 eq.) was added. The mixture was stirred at 80 °C for 12 h. Water (30 mL) was added to the mixture, and the resulting mixture was extracted with ethyl acetate (20 mL x 3). The combined organic phase was washed with brine (20 mL), dried over sodium sulfate, filtered, and concentrated in vacuum. The residue was purified by silica gel chromatography (dichloromethane:methanol = 1:0 to 20:1). Tert-butyl 4-[4-[4-(3-amino-6-chloro-pyridazin-4-yl)pyrazol-1-yl]but-2-ynyl]piperazine-1-carboxylate (600 mg, 1.34 mmol, 71% yield) was obtained as a brown solid.
工程8
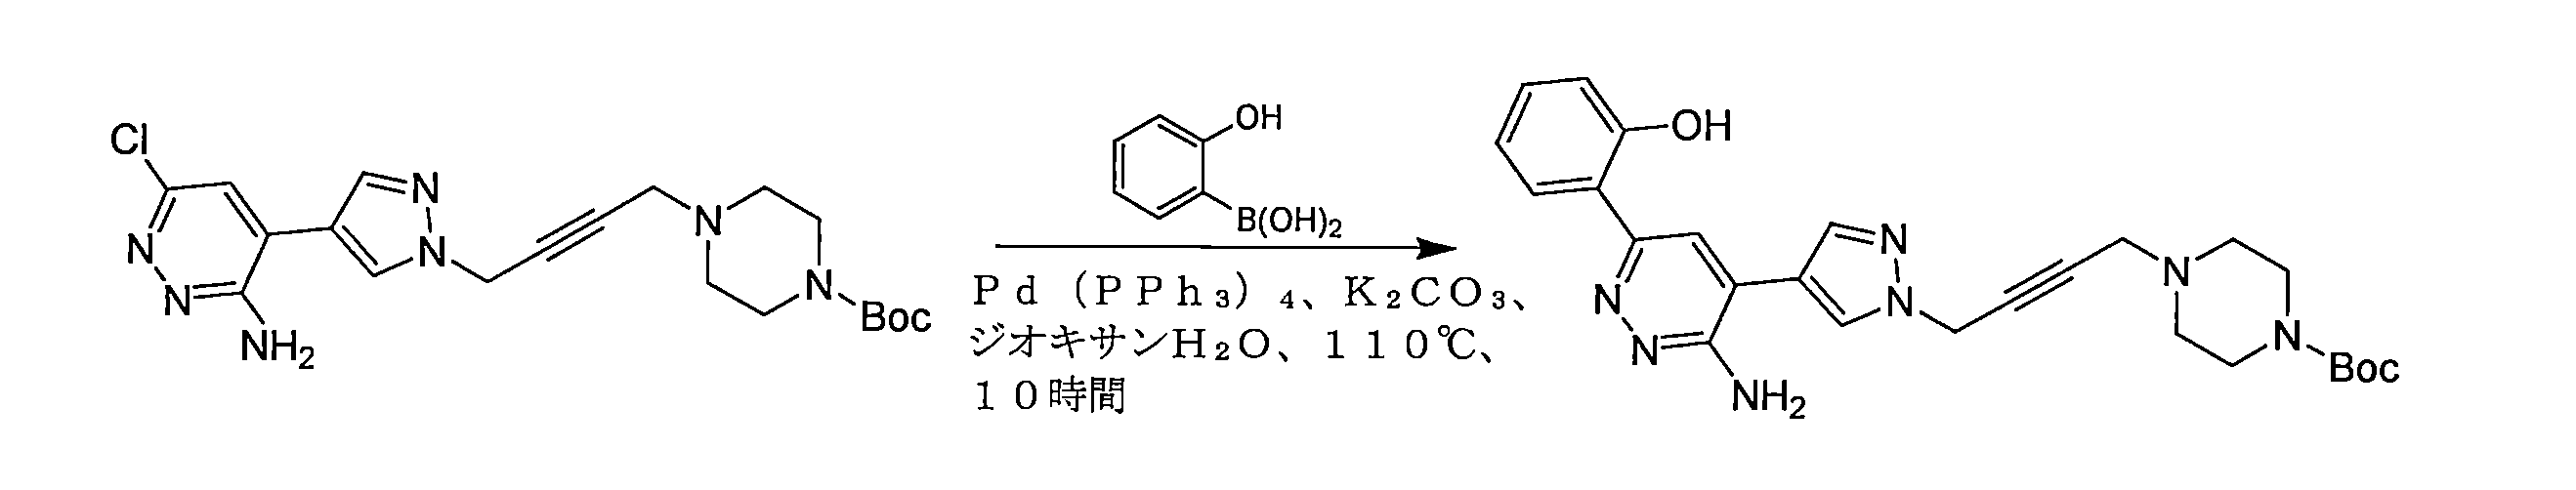
ジオキサン(10mL)および水(2mL)中のtert-ブチル4-[4-[4-(3-アミノ-6-クロロ-ピリダジン-4-イル)ピラゾール-1-イル]ブタ-2-イニル]ピペラジン-1-カルボキシレート(500mg、1.16mmol、1当量)、(2-ヒドロキシフェニル)ボロン酸(192mg、1.39mmol、1.2当量)の溶液に、炭酸カリウム(400mg、2.89mmol、2.5当量)およびテトラキス[トリフェニルホスフィン]パラジウム(0)(134mg、0.12mmol、0.1当量)を窒素下で添加した。反応混合物を110℃で10時間撹拌した。水(100mL)を混合物に加え、得られた混合物を酢酸エチル(50mL×3)で抽出した。合わせた有機相をブライン(50mL)で洗浄し、硫酸ナトリウムで乾燥させ、濾過し、真空中で濃縮した。残渣を分取薄層クロマトグラフィー(ジクロロメタン:メタノール=10:1)によって精製した。Tert-ブチル4-[4-[4-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]ピラゾール-1-イル]ブタ-2-イニル]ピペラジン-1-カルボキシレート(340mg、0.65mmol、収率56%)を淡い黄色の固体として得た。 To a solution of tert-butyl 4-[4-[4-(3-amino-6-chloro-pyridazin-4-yl)pyrazol-1-yl]but-2-ynyl]piperazine-1-carboxylate (500 mg, 1.16 mmol, 1 eq.), (2-hydroxyphenyl)boronic acid (192 mg, 1.39 mmol, 1.2 eq.) in dioxane (10 mL) and water (2 mL), potassium carbonate (400 mg, 2.89 mmol, 2.5 eq.) and tetrakis[triphenylphosphine]palladium(0) (134 mg, 0.12 mmol, 0.1 eq.) were added under nitrogen. The reaction mixture was stirred at 110 °C for 10 h. Water (100 mL) was added to the mixture and the resulting mixture was extracted with ethyl acetate (50 mL × 3). The combined organic phase was washed with brine (50 mL), dried over sodium sulfate, filtered and concentrated in vacuo. The residue was purified by preparative thin-layer chromatography (dichloromethane:methanol=10:1). Tert-butyl 4-[4-[4-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]pyrazol-1-yl]but-2-ynyl]piperazine-1-carboxylate (340 mg, 0.65 mmol, 56% yield) was obtained as a pale yellow solid.
上記の他の実施例について記載される手順を使用して、以下のスキームに従って、tert-ブチル4-[4-[4-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]ピラゾール-1-イル]ブタ-2-イニル]ピペラジン-1-カルボキシレートを最終化合物に変換した。

例示的な化合物83の例示的な合成

工程1

250mLの丸底フラスコに、DCM(100mL)中のメチル4-(1-ヒドロキシエチル)ベンゾエート(2.5g、13.87mmol、1当量)、トリフェニルホスフィンジブロミド(5.9g、13.87mmol、1当量)を入れた。得られた混合物を室温で3時間撹拌した。反応混合物を減圧下で濃縮し、残渣を、酢酸エチル/石油エーテル(v:v=1:12)で溶出するシリカゲルカラム上に適用した。これにより、2.3g(68%)のメチル4-(1-ブロモエチル)ベンゾエートを無色の油として得た。 A 250 mL round bottom flask was charged with methyl 4-(1-hydroxyethyl)benzoate (2.5 g, 13.87 mmol, 1 equiv.), triphenylphosphine dibromide (5.9 g, 13.87 mmol, 1 equiv.) in DCM (100 mL). The resulting mixture was stirred at room temperature for 3 h. The reaction mixture was concentrated under reduced pressure and the residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (v:v=1:12). This gave 2.3 g (68%) of methyl 4-(1-bromoethyl)benzoate as a colorless oil.
工程2

30mLのバイアルに、DMF(15mL)中のメチル4-(1-ブロモエチル)ベンゾエート(1.2g、4.94mmol、1当量)、6-[2-(メトキシメトキシ)フェニル]-4-(1H-ピラゾール-4-イル)ピリダジン-3-アミン(1.2g、3.95mmol、0.80当量)、K2CO3(2.0g、14.81mmol、3.00当量)、NaI(739.9mg、4.94mmol、1.00当量)を入れた。得られた混合物を、油浴中で、60℃で3時間撹拌した。次いで、反応混合物を水(50mL)で希釈し、ジクロロメタン(50mL×3)で抽出した。合わせた有機層を水(50mL×2)およびブライン(50mL)で洗浄し、無水硫酸ナトリウムで乾燥させ、減圧下で濃縮した。残渣を、ジクロロメタン/メタノール(v:v=10:1)で溶出するシリカゲルカラム上に適用した。これにより、600mg(26%)のメチル4-[1-(4-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-1H-ピラゾール-1-イル)エチル]ベンゾエートを黄色の油として得た。 A 30 mL vial was charged with methyl 4-(1-bromoethyl)benzoate (1.2 g, 4.94 mmol, 1 equiv.), 6-[2-(methoxymethoxy)phenyl]-4-(1H-pyrazol-4-yl)pyridazin-3-amine (1.2 g, 3.95 mmol, 0.80 equiv.), K2CO3 (2.0 g, 14.81 mmol, 3.00 equiv.), NaI (739.9 mg, 4.94 mmol, 1.00 equiv.) in DMF (15 mL). The resulting mixture was stirred in an oil bath at 60 °C for 3 h. The reaction mixture was then diluted with water (50 mL) and extracted with dichloromethane (50 mL x 3). The combined organic layers were washed with water (50 mL x 2) and brine (50 mL), dried over anhydrous sodium sulfate, and concentrated under reduced pressure. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (v:v=10:1). This gave 600 mg (26%) of methyl 4-[1-(4-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-1H-pyrazol-1-yl)ethyl]benzoate as a yellow oil.
工程3

100mLの丸底フラスコに、THF(20mL)中のメチル4-[1-(4-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-1H-ピラゾール-1-イル)エチル]ベンゾエート(600mg、1.31mmol、1当量)の溶液を入れ、それにLiAlH4(495.6mg、13.06mmol、10.00当量)を0℃で添加した。得られた混合物を室温で3時間撹拌した。次いで、反応物を、Na2SO4.10H2O(4.8g、15.0mmol)を添加することによってクエンチした。得られた混合物を室温で3時間撹拌した。反応混合物中の固形分を濾過し、濾液を減圧下で濃縮した。残渣を、ジクロロメタン/メタノール(v:v=10:1)で溶出するシリカゲルカラム上に適用した。これにより、310mg(55%)の[4-[1-(4-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-1H-ピラゾール-1-イル)エチル]フェニル]メタノールを黄色の油として得た。 A 100 mL round bottom flask was charged with a solution of methyl 4-[1-(4-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-1H-pyrazol-1-yl)ethyl]benzoate (600 mg, 1.31 mmol, 1 equiv.) in THF (20 mL), to which LiAlH4 (495.6 mg, 13.06 mmol, 10.00 equiv.) was added at 0°C. The resulting mixture was stirred at room temperature for 3 hours. The reaction was then quenched by adding Na2SO4.10H2O (4.8 g, 15.0 mmol). The resulting mixture was stirred at room temperature for 3 hours. The solids in the reaction mixture were filtered and the filtrate was concentrated under reduced pressure. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (v:v=10:1). This gave 310 mg (55%) of [4-[1-(4-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-1H-pyrazol-1-yl)ethyl]phenyl]methanol as a yellow oil.
工程4
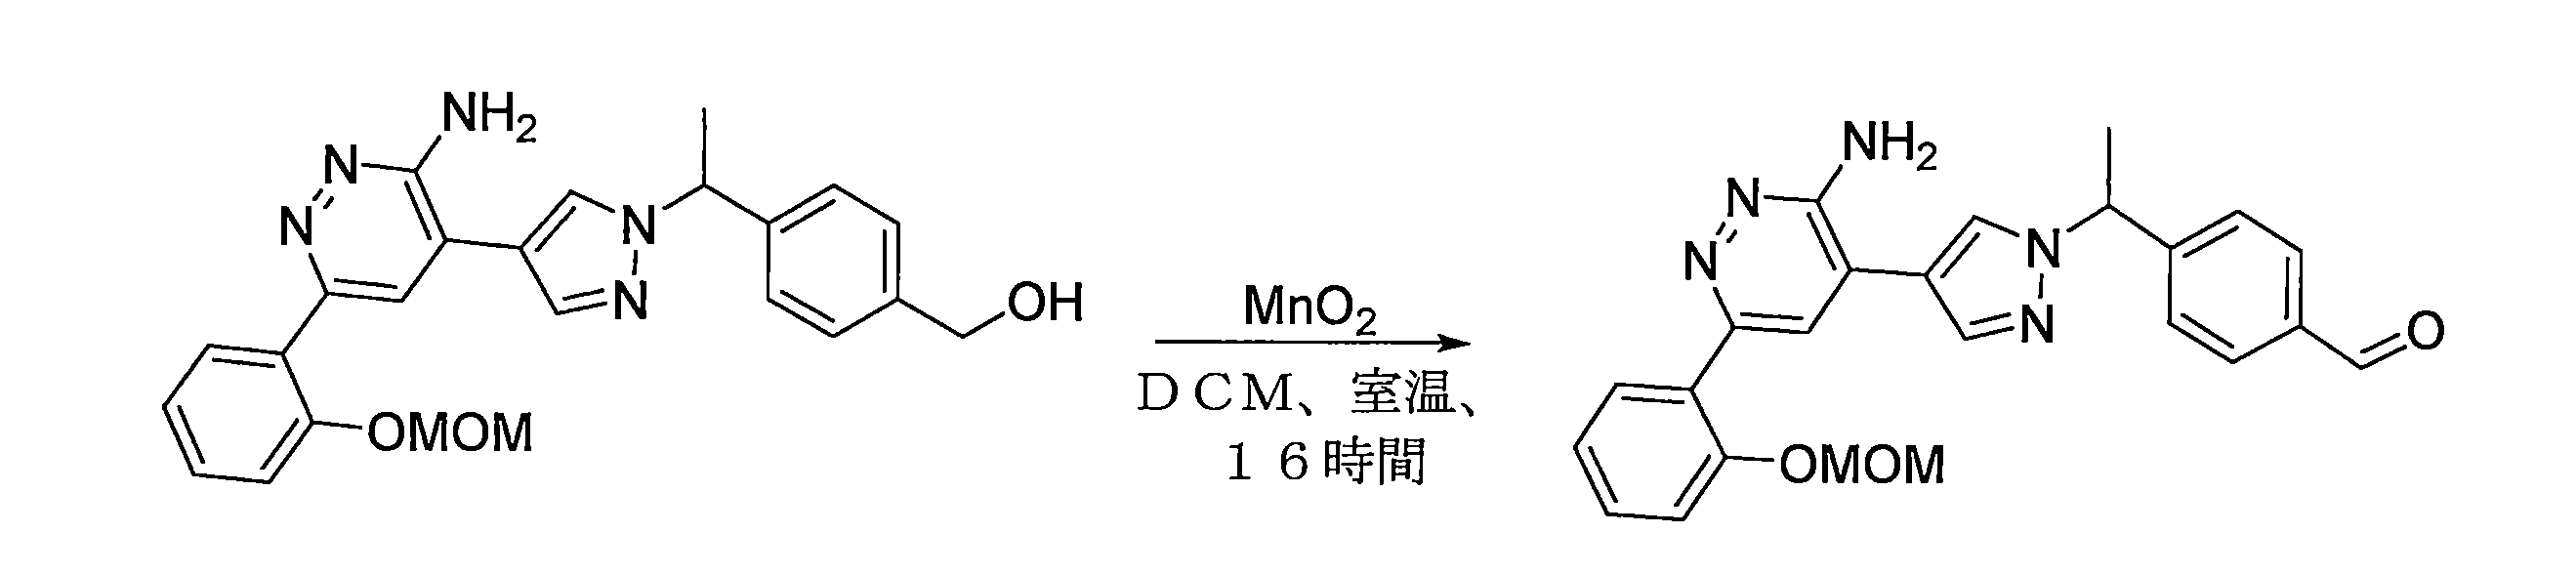
100mLの丸底フラスコに、DCM(10.0mL、117mmol、218当量)中の[4-[1-(4-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-1H-ピラゾール-1-イル)エチル]フェニル]メタノール(310mg、0.72mmol、1当量)の溶液を入れ、それに二酸化マンガン(1.2g、14.37mmol、20当量)を室温で添加した。得られた混合物を、室温で16時間撹拌した。固形分を濾過し、濾液を減圧下で濃縮した。これにより、286mg(93%)の4-[1-(4-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-1H-ピラゾール-1-イル)エチル]ベンズアルデヒドを黄色の固体として得た。 A 100 mL round bottom flask was charged with a solution of [4-[1-(4-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-1H-pyrazol-1-yl)ethyl]phenyl]methanol (310 mg, 0.72 mmol, 1 equiv.) in DCM (10.0 mL, 117 mmol, 218 equiv.) to which manganese dioxide (1.2 g, 14.37 mmol, 20 equiv.) was added at room temperature. The resulting mixture was stirred at room temperature for 16 hours. The solids were filtered and the filtrate was concentrated under reduced pressure. This afforded 286 mg (93%) of 4-[1-(4-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-1H-pyrazol-1-yl)ethyl]benzaldehyde as a yellow solid.
工程5

100mLの丸底フラスコに、DCM(10mL)中の4-[1-(4-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-1H-ピラゾール-1-イル)エチル]ベンズアルデヒド(300.0mg、0.70mmol、1当量)、ベンジルピペラジン-1-カルボキシレート(185mg、0.84mmol、1.2当量)、HOAc(0.1mL、1.75mmol、2.50当量)を入れた。得られた混合物を0.5時間撹拌し、次いで、NaBH(OAc)3(444.1mg、2.10mmol、3当量)を添加した。得られた混合物を室温でさらに16時間撹拌した。次いで、反応混合物を水(20mL)によってクエンチし、ジクロロメタン(20mL×3)で抽出した。合わせた有機層を水(20mL×2)およびブライン(20mL)で洗浄し、無水硫酸ナトリウムで乾燥させ、減圧下で濃縮した。残渣を、ジクロロメタン/メタノール(v:v=10:1)で溶出するシリカゲルカラム上に適用した。これにより、260mg(59%)のベンジル4-([4-[1-(4-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-1H-ピラゾール-1-イル)エチル]フェニル]メチル)ピペラジン-1-カルボキシレートを黄色の油として得た。 A 100 mL round bottom flask was charged with 4-[1-(4-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-1H-pyrazol-1-yl)ethyl]benzaldehyde (300.0 mg, 0.70 mmol, 1 eq.), benzyl piperazine-1-carboxylate (185 mg, 0.84 mmol, 1.2 eq.), HOAc (0.1 mL, 1.75 mmol, 2.50 eq.) in DCM (10 mL). The resulting mixture was stirred for 0.5 h and then NaBH(OAc)3 (444.1 mg, 2.10 mmol, 3 eq.) was added. The resulting mixture was stirred for an additional 16 h at room temperature. The reaction mixture was then quenched by water (20 mL) and extracted with dichloromethane (20 mL x 3). The combined organic layers were washed with water (20 mL x 2) and brine (20 mL), dried over anhydrous sodium sulfate, and concentrated under reduced pressure. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (v:v=10:1). This gave 260 mg (59%) of benzyl 4-([4-[1-(4-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-1H-pyrazol-1-yl)ethyl]phenyl]methyl)piperazine-1-carboxylate as a yellow oil.
工程6

窒素不活性雰囲気下でパージし、維持した100mLの丸底フラスコに、THF(8mL)中のベンジル4-([4-[1-(4-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-1H-ピラゾール-1-イル)エチル]フェニル]メチル)ピペラジン-1-カルボキシレート(260.0mg、0.41mmol、1当量)の溶液を入れ、それに水素化トリエチルホウ素リチウム(2.1mL、2.05mmol、5.00当量)を0℃で添加した。反応混合物を室温で16時間撹拌した。次いで、反応混合物を水(30mL)によってクエンチし、ジクロロメタン(30mL×3)で抽出した。合わせた有機層をブライン(30mL)で洗浄し、無水ナトリウムで乾燥させ、濃縮した。これにより、200mg(98%)の6-[2-(メトキシメトキシ)フェニル]-4-[1-(1-[4-[(ピペラジン-1-イル)メチル]フェニル]エチル)-1H-ピラゾール-4-イル]ピリダジン-3-アミンを黄色の油として得た。 A 100 mL round bottom flask purged and maintained under nitrogen inert atmosphere was charged with a solution of benzyl 4-([4-[1-(4-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-1H-pyrazol-1-yl)ethyl]phenyl]methyl)piperazine-1-carboxylate (260.0 mg, 0.41 mmol, 1 equiv.) in THF (8 mL) to which lithium triethylborohydride (2.1 mL, 2.05 mmol, 5.00 equiv.) was added at 0°C. The reaction mixture was stirred at room temperature for 16 hours. The reaction mixture was then quenched by water (30 mL) and extracted with dichloromethane (30 mL x 3). The combined organic layers were washed with brine (30 mL), dried over anhydrous sodium, and concentrated. This gave 200 mg (98%) of 6-[2-(methoxymethoxy)phenyl]-4-[1-(1-[4-[(piperazin-1-yl)methyl]phenyl]ethyl)-1H-pyrazol-4-yl]pyridazin-3-amine as a yellow oil.
他の実施例について上に記載される手順を使用して、以下のスキームに従って、6-[2-(メトキシメトキシ)フェニル]-4-[1-(1-[4-[(ピペラジン-1-イル)メチル]フェニル]エチル)-1H-ピラゾール-4-イル]ピリダジン-3-アミンを表題化合物に変換した。
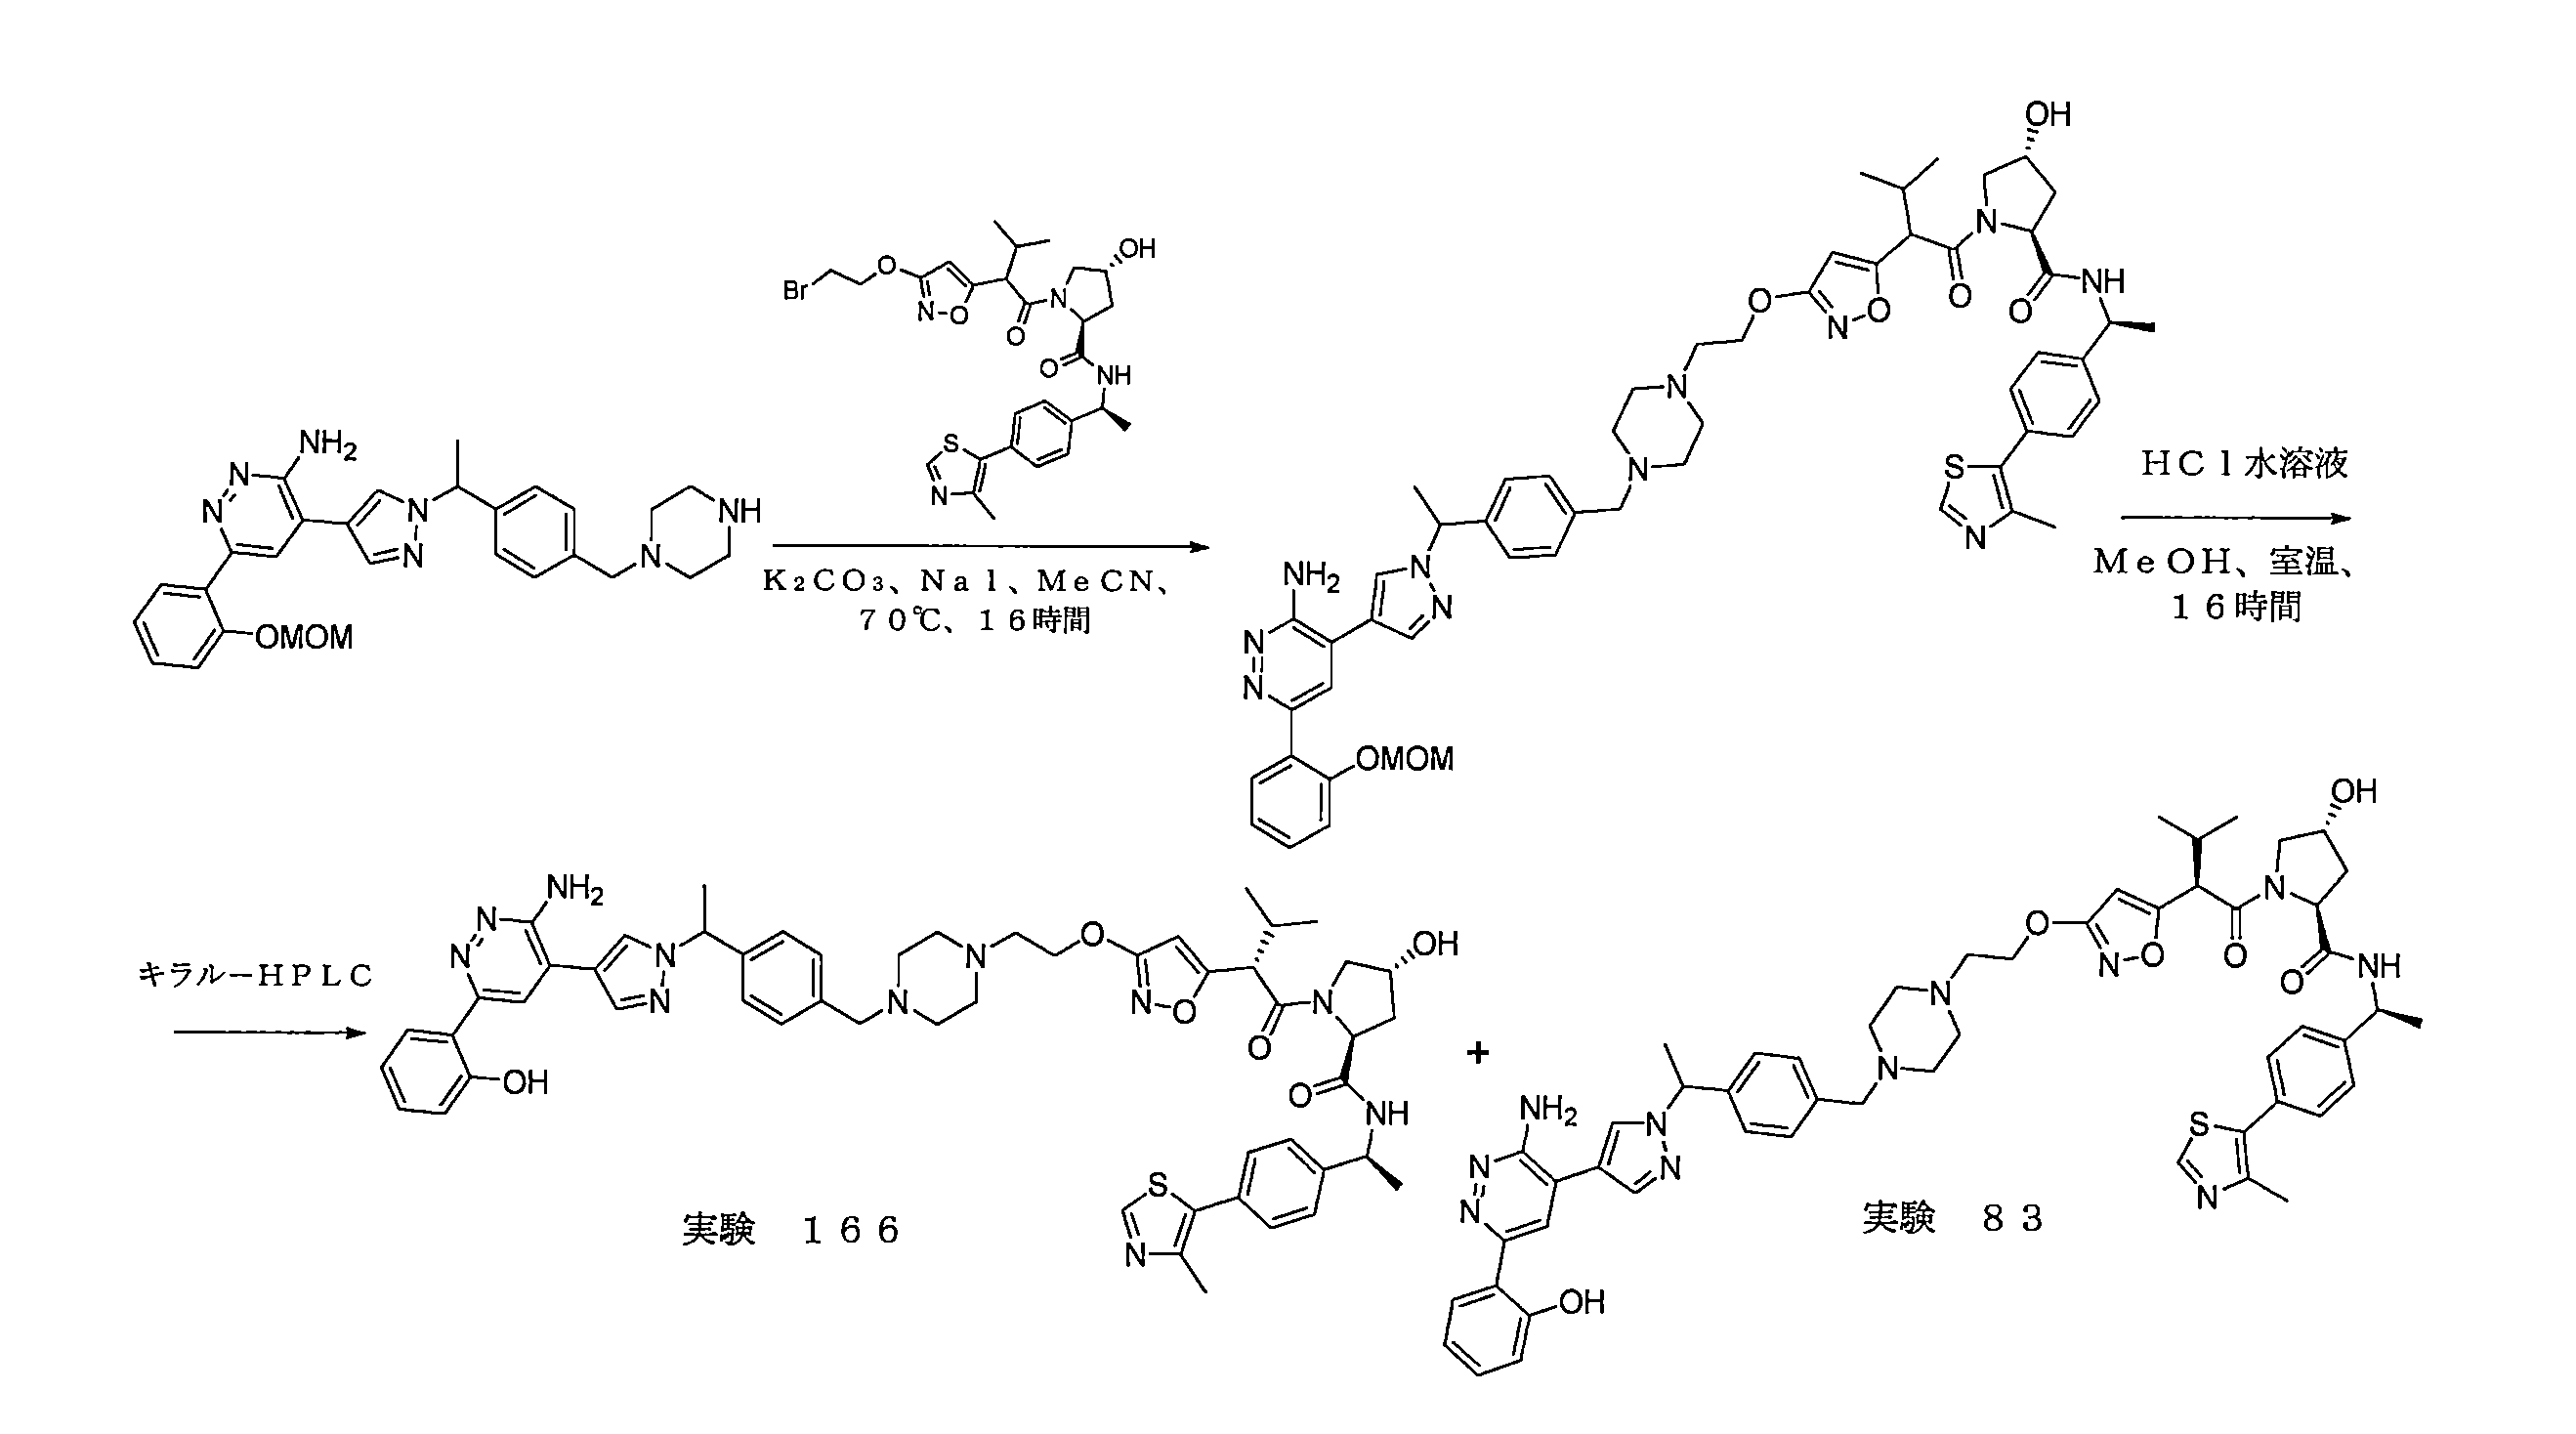
例示的な化合物167の例示的な合成

工程1

ジオキサン(100mL)中のtert-ブチルN-[(1S)-1-(4-ブロモフェニル)エチル]カルバメート(5g、16.7mmol、1当量)および4,4,5,5-テトラメチル-2-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)-1,3,2-ジオキサボロラン(5.08g、20.0mmol、1.2当量)の溶液に、[1,1’-ビス(ジフェニルホスフィノ)フェロセン]ジクロロパラジウム(II)(975mg、1.33mmol、0.08当量)および酢酸カリウム(3.27g、33.31mmol、2当量)を添加した。混合物を90℃で2時間撹拌した。反応混合物を濃縮し、粗生成物を得た。粗生成物を、シリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=1:0~10:1)により精製した。化合物tert-ブチルN-[(1S)-1-[4-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)フェニル]エチル]カルバメート(8.2g、16.44mmol、収率99%)を黄色の油として得た。 To a solution of tert-butyl N-[(1S)-1-(4-bromophenyl)ethyl]carbamate (5 g, 16.7 mmol, 1 eq.) and 4,4,5,5-tetramethyl-2-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)-1,3,2-dioxaborolane (5.08 g, 20.0 mmol, 1.2 eq.) in dioxane (100 mL), [1,1'-bis(diphenylphosphino)ferrocene]dichloropalladium(II) (975 mg, 1.33 mmol, 0.08 eq.) and potassium acetate (3.27 g, 33.31 mmol, 2 eq.) were added. The mixture was stirred at 90 °C for 2 h. The reaction mixture was concentrated to give the crude product. The crude product was purified by silica gel chromatography (petroleum ether:ethyl acetate = 1:0 to 10:1). The compound tert-butyl N-[(1S)-1-[4-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)phenyl]ethyl]carbamate (8.2 g, 16.44 mmol, 99% yield) was obtained as a yellow oil.
工程2

ジオキサン(130mL)および水(22mL)の混合物中のエチル5-ブロモチアゾール-4-カルボキシレート(3.49g、14.77mmol、0.9当量)、tert-ブチルN-[(1S)-1-[4-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)フェニル]エチル]カルバメート(5.70g、16.41mmol、1当量)、[1,1’-ビス(ジフェニルホスフィノ)フェロセン]ジクロロパラジウム(II)(120mg、0.16mmol、0.01当量)、炭酸カリウム(4.54g、32.83mmol、2当量)の混合物を、脱気し、窒素で3回パージし、次いで、混合物を窒素雰囲気下で、80℃で10時間撹拌した。反応混合物を濃縮し、水(50mL)を添加し、次いで、酢酸エチル(60mL)で2回抽出した。合わせた有機層を、塩化ナトリウムの飽和水溶液(50mL)で2回洗浄し、無水硫酸ナトリウム上で乾燥させ、濃縮して粗生成物を得た。粗生成物を、シリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=1:0~1:1)により精製した。化合物エチル5-[4-[(1S)-1-(tert-ブトキシカルボニルアミノ)エチル]フェニル]チアゾール-4-カルボキシレート(5.07g、13.48mmol、収率82%)を黄色の固体として得た。 A mixture of ethyl 5-bromothiazole-4-carboxylate (3.49 g, 14.77 mmol, 0.9 eq.), tert-butyl N-[(1S)-1-[4-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)phenyl]ethyl]carbamate (5.70 g, 16.41 mmol, 1 eq.), [1,1'-bis(diphenylphosphino)ferrocene]dichloropalladium(II) (120 mg, 0.16 mmol, 0.01 eq.), potassium carbonate (4.54 g, 32.83 mmol, 2 eq.) in a mixture of dioxane (130 mL) and water (22 mL) was degassed and purged with nitrogen three times, then the mixture was stirred at 80 °C under nitrogen atmosphere for 10 h. The reaction mixture was concentrated, water (50 mL) was added, and then extracted twice with ethyl acetate (60 mL). The combined organic layers were washed twice with a saturated aqueous solution of sodium chloride (50 mL), dried over anhydrous sodium sulfate, and concentrated to obtain the crude product. The crude product was purified by silica gel chromatography (petroleum ether:ethyl acetate=1:0 to 1:1). The compound ethyl 5-[4-[(1S)-1-(tert-butoxycarbonylamino)ethyl]phenyl]thiazole-4-carboxylate (5.07 g, 13.48 mmol, yield 82%) was obtained as a yellow solid.
工程3

テトラヒドロフラン(60mL)中のエチル5-[4-[(1S)-1-(tert-ブトキシカルボニルアミノ)エチル]フェニル]チアゾール-4-カルボキシレート(4g、10.63mmol、1当量)の溶液を、窒素雰囲気下で-78℃に冷却した。次いで、水素化ジイソブチルアルミニウムTHF溶液(1M、11.69mL、1.1当量)を-78℃で、滴下で添加した。反応混合物を-78℃で0.5時間撹拌した。メタノール(2mL)を添加して反応物にクエンチし、続いて水(50mL)を添加した。混合物を酢酸エチル(40mL×2)で抽出した。有機層を硫酸ナトリウムで乾燥させ、次いで、真空下で濃縮して、残渣を得た。残渣を、シリカゲルカラムクロマトグラフィー(石油エーテル中10~50%酢酸エチル)によって精製した。化合物tert-ブチルN-[(1S)-1-[4-(4-ホルミルチアゾール-5-イル)フェニル]エチル]カルバメート(3.2g、9.63mmol、収率91%)を白色の固体として得た。 A solution of ethyl 5-[4-[(1S)-1-(tert-butoxycarbonylamino)ethyl]phenyl]thiazole-4-carboxylate (4 g, 10.63 mmol, 1 equiv.) in tetrahydrofuran (60 mL) was cooled to −78°C under nitrogen atmosphere. Then, diisobutylaluminum hydride THF solution (1 M, 11.69 mL, 1.1 equiv.) was added dropwise at −78°C. The reaction mixture was stirred at −78°C for 0.5 h. The reaction was quenched by adding methanol (2 mL) followed by water (50 mL). The mixture was extracted with ethyl acetate (40 mL x 2). The organic layer was dried over sodium sulfate and then concentrated under vacuum to give a residue. The residue was purified by silica gel column chromatography (10-50% ethyl acetate in petroleum ether). The compound tert-butyl N-[(1S)-1-[4-(4-formylthiazol-5-yl)phenyl]ethyl]carbamate (3.2 g, 9.63 mmol, 91% yield) was obtained as a white solid.
工程4
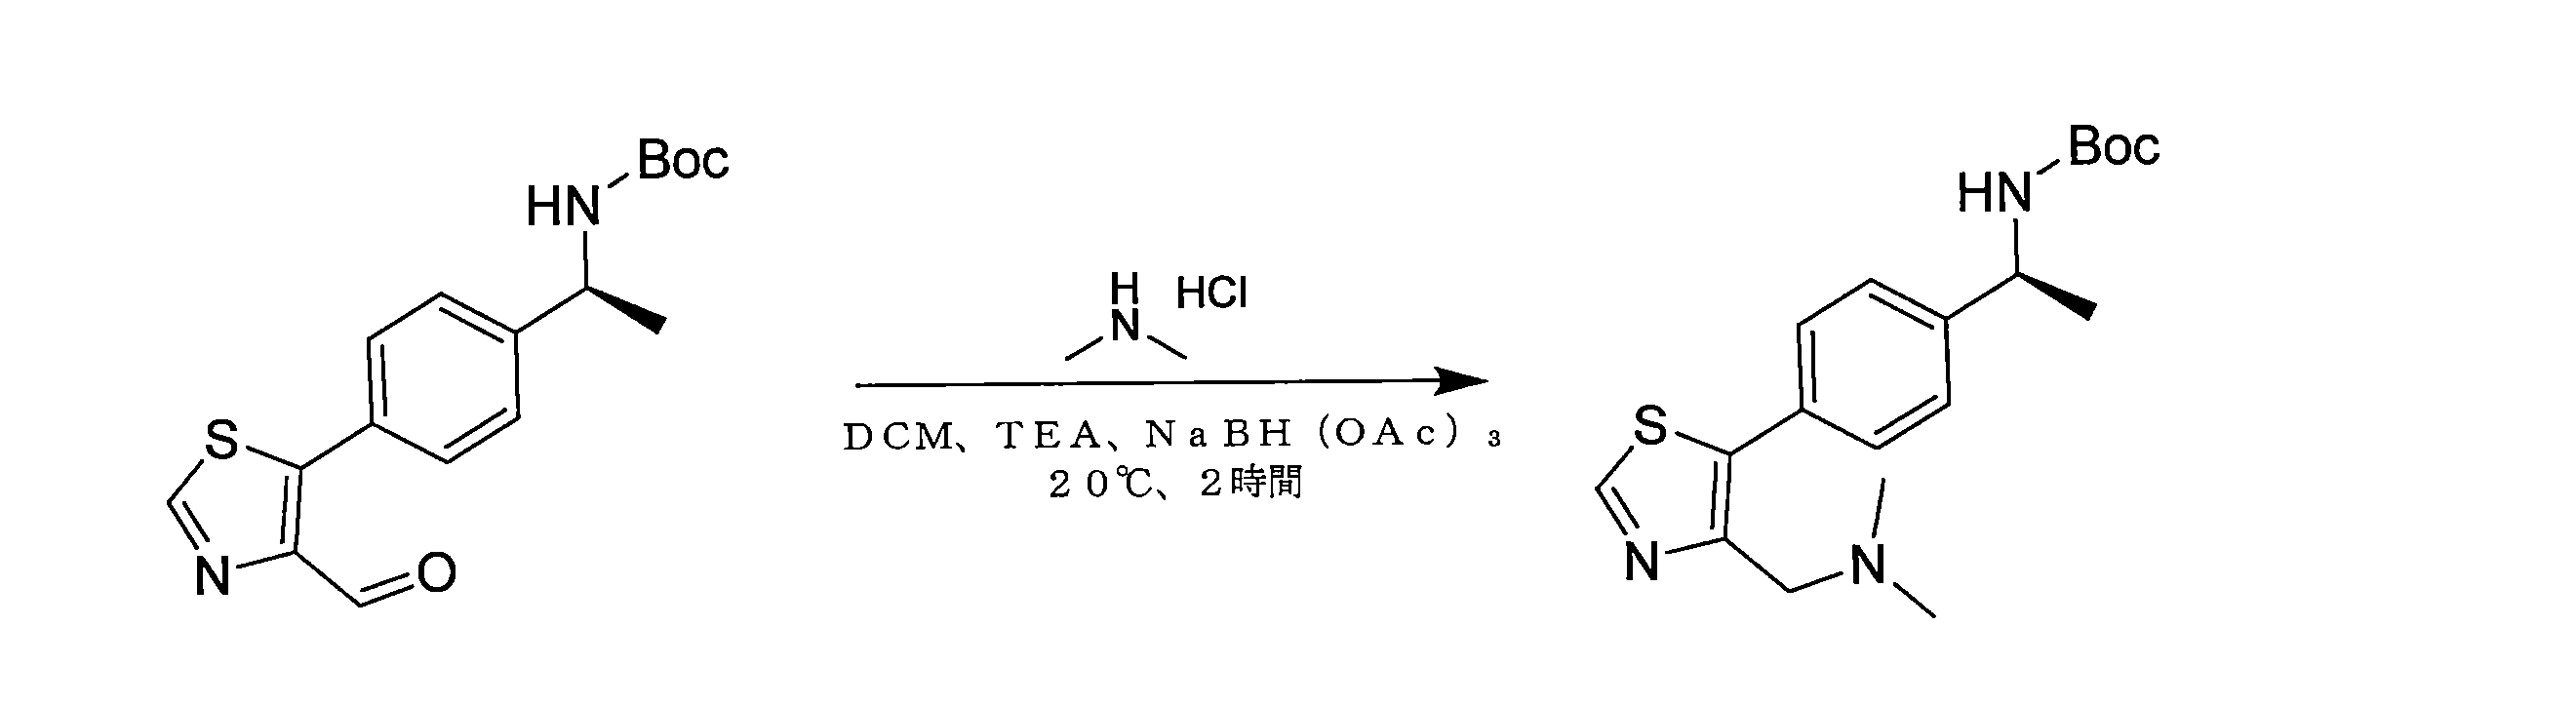
ジクロロメタン(30mL)中のジメチルアミン塩酸塩(368mg、4.51mmol、0.41mL、3当量)の溶液に、トリエチルアミン(609mg、6.02mmol、0.84mL、4当量)、ジクロロメタン(10mL)中のtert-ブチルN-[(1S)-1-[4-(4-ホルミルチアゾール-5-イル)フェニル]エチル]カルバメート(500mg、1.50mmol、1当量)の溶液、およびトリアセトキシホウ素水素化ナトリウム(1.28g、6.02mmol、4当量)を添加し、混合物を20℃で2時間撹拌した。反応混合物を、水(50mL)を添加することによってクエンチし、ジクロロメタン(40mL)で2回抽出した。合わせた有機層を飽和塩化ナトリウム水溶液(60mL)で2回洗浄し、無水硫酸ナトリウムで乾燥させ、濃縮して、生成物を得た。粗生成物をシリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=1:0~0:1)によって精製した。化合物tert-ブチルN-[(1S)-1-[4-[4-[(ジメチルアミノ)メチル]チアゾール-5-イル]フェニル]エチル]カルバメート(364mg、1.01mmol、収率67%)を黄色の油として得た。 To a solution of dimethylamine hydrochloride (368 mg, 4.51 mmol, 0.41 mL, 3 eq.) in dichloromethane (30 mL), triethylamine (609 mg, 6.02 mmol, 0.84 mL, 4 eq.), a solution of tert-butyl N-[(1S)-1-[4-(4-formylthiazol-5-yl)phenyl]ethyl]carbamate (500 mg, 1.50 mmol, 1 eq.) in dichloromethane (10 mL), and sodium triacetoxyborohydride (1.28 g, 6.02 mmol, 4 eq.) were added and the mixture was stirred at 20 °C for 2 h. The reaction mixture was quenched by adding water (50 mL) and extracted twice with dichloromethane (40 mL). The combined organic layers were washed twice with saturated aqueous sodium chloride solution (60 mL), dried over anhydrous sodium sulfate, and concentrated to give the product. The crude product was purified by silica gel chromatography (petroleum ether:ethyl acetate = 1:0 to 0:1). The compound tert-butyl N-[(1S)-1-[4-[4-[(dimethylamino)methyl]thiazol-5-yl]phenyl]ethyl]carbamate (364 mg, 1.01 mmol, yield 67%) was obtained as a yellow oil.
例示的な化合物76について記載される手順を使用して、tert-ブチルN-[(1S)-1-[4-[4-[(ジメチルアミノ)メチル]チアゾール-5-イル]フェニル]エチル]カルバメートを最終生成物に変換した。 Using the procedure described for exemplary compound 76, tert-butyl N-[(1S)-1-[4-[4-[(dimethylamino)methyl]thiazol-5-yl]phenyl]ethyl]carbamate was converted to the final product.
例示的な化合物168の例示的な合成

工程1

水(40mL)およびテトラヒドロフラン(20mL)中のエチル5-[4-[(1S)-1-(tert-ブトキシカルボニルアミノ)エチル]フェニル]チアゾール-4-カルボキシレート(900mg、2.39mmol、1当量)および塩化カルシウム(796mg、7.17mmol、3当量)の混合物に、水素化ホウ素ナトリウム(181mg、4.78mmol、2当量)を窒素下で、0℃で添加した。混合物を5℃に温め、2時間撹拌した。塩酸(5M、4.50mL)およびアセトン(4.95mL)を0℃で添加し、反応混合物を5℃でさらに0.5時間撹拌した。5M水酸化ナトリウムを用いてpHを5に調整した。水層を、酢酸エチル(20mL×4)で抽出した。合わせた有機層をブライン(20mL×2)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、真空下で濃縮させた。残渣を半分取逆相HPLC(基本条件、カラム:Waters Xbridge 150*50 10um;移動相:[水(0.05%水酸化アンモニアv/v)-ACN];B%:25%~55%、11.5分)によって精製した。化合物tert-ブチルN-[(1S)-1-[4-[4-(ヒドロキシメチル)チアゾール-5-イル]フェニル]エチル]カルバメート(400mg、1.20mmol、収率50%)を白色の固体として得た。 To a mixture of ethyl 5-[4-[(1S)-1-(tert-butoxycarbonylamino)ethyl]phenyl]thiazole-4-carboxylate (900 mg, 2.39 mmol, 1 eq.) and calcium chloride (796 mg, 7.17 mmol, 3 eq.) in water (40 mL) and tetrahydrofuran (20 mL), sodium borohydride (181 mg, 4.78 mmol, 2 eq.) was added under nitrogen at 0°C. The mixture was warmed to 5°C and stirred for 2 h. Hydrochloric acid (5 M, 4.50 mL) and acetone (4.95 mL) were added at 0°C and the reaction mixture was stirred at 5°C for an additional 0.5 h. The pH was adjusted to 5 with 5 M sodium hydroxide. The aqueous layer was extracted with ethyl acetate (20 mL x 4). The combined organic layers were washed with brine (20 mL x 2), dried over anhydrous sodium sulfate, filtered and concentrated under vacuum. The residue was purified by semi-preparative reverse phase HPLC (basic conditions, column: Waters Xbridge 150*50 10um; mobile phase: [water (0.05% ammonia hydroxide v/v)-ACN]; B%: 25%-55%, 11.5 min). The compound tert-butyl N-[(1S)-1-[4-[4-(hydroxymethyl)thiazol-5-yl]phenyl]ethyl]carbamate (400mg, 1.20mmol, 50% yield) was obtained as a white solid.
例示的な化合物76について記載される手順を使用して、tert-ブチルN-[(1S)-1-[4-[4-(ヒドロキシメチル)チアゾール-5-イル]フェニル]エチル]カルバメートを最終生成物に変換した。 Using the procedure described for exemplary compound 76, tert-butyl N-[(1S)-1-[4-[4-(hydroxymethyl)thiazol-5-yl]phenyl]ethyl]carbamate was converted to the final product.
例示的な化合物211および219も、上に記載されるものと類似の手順を使用して調製する。 Exemplary compounds 211 and 219 are also prepared using procedures similar to those described above.
例示的な化合物169の例示的な合成

工程1

例示的な化合物168について記載される手順を使用して調製した。 Prepared using the procedure described for exemplary compound 168.
工程2
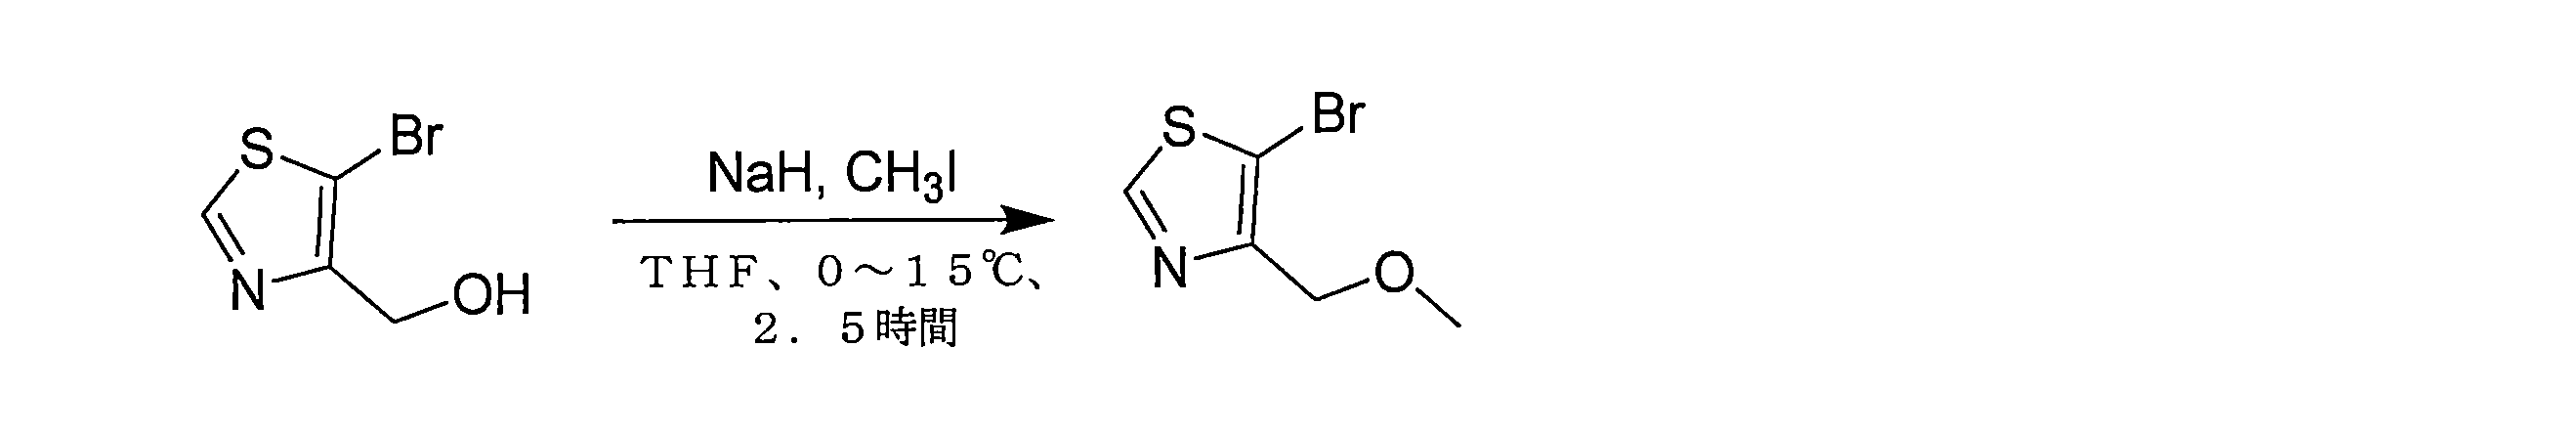
THF(30mL)中の(5-ブロモチアゾール-4-イル)メタノール(980mg、5.05mmol、1当量)の溶液に、水素化ナトリウム(404mg、10.10mmol、2当量)を窒素下で、0℃で添加した。混合物を0℃で30分間撹拌し、次いで、ヨードメタン(3.58g、25.25mmol、1.57mL、5当量)を0℃で添加した。混合物を窒素下で、15℃において2時間撹拌した。混合物を0℃に冷却し、塩化アンモニウムの飽和水溶液によってゆっくりクエンチし、次いで、酢酸エチル(50mL×3)で抽出した。合わせた有機相をブライン(25mL×2)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、真空下で濃縮した。残渣を、シリカゲルクロマトグラフィー(石油エーテル/酢酸エチル=20/1~10/1)によって精製し、5-ブロモ-4-(メトキシメチル)チアゾール(910mg、4.37mmol、収率86%)を黄色の固体として得た。 To a solution of (5-bromothiazol-4-yl)methanol (980 mg, 5.05 mmol, 1 eq.) in THF (30 mL) was added sodium hydride (404 mg, 10.10 mmol, 2 eq.) under nitrogen at 0°C. The mixture was stirred at 0°C for 30 minutes, then iodomethane (3.58 g, 25.25 mmol, 1.57 mL, 5 eq.) was added at 0°C. The mixture was stirred under nitrogen at 15°C for 2 hours. The mixture was cooled to 0°C and slowly quenched with a saturated aqueous solution of ammonium chloride, then extracted with ethyl acetate (50 mL x 3). The combined organic phase was washed with brine (25 mL x 2), dried over anhydrous sodium sulfate, filtered, and concentrated under vacuum. The residue was purified by silica gel chromatography (petroleum ether/ethyl acetate = 20/1 to 10/1) to obtain 5-bromo-4-(methoxymethyl)thiazole (910 mg, 4.37 mmol, yield 86%) as a yellow solid.
工程3
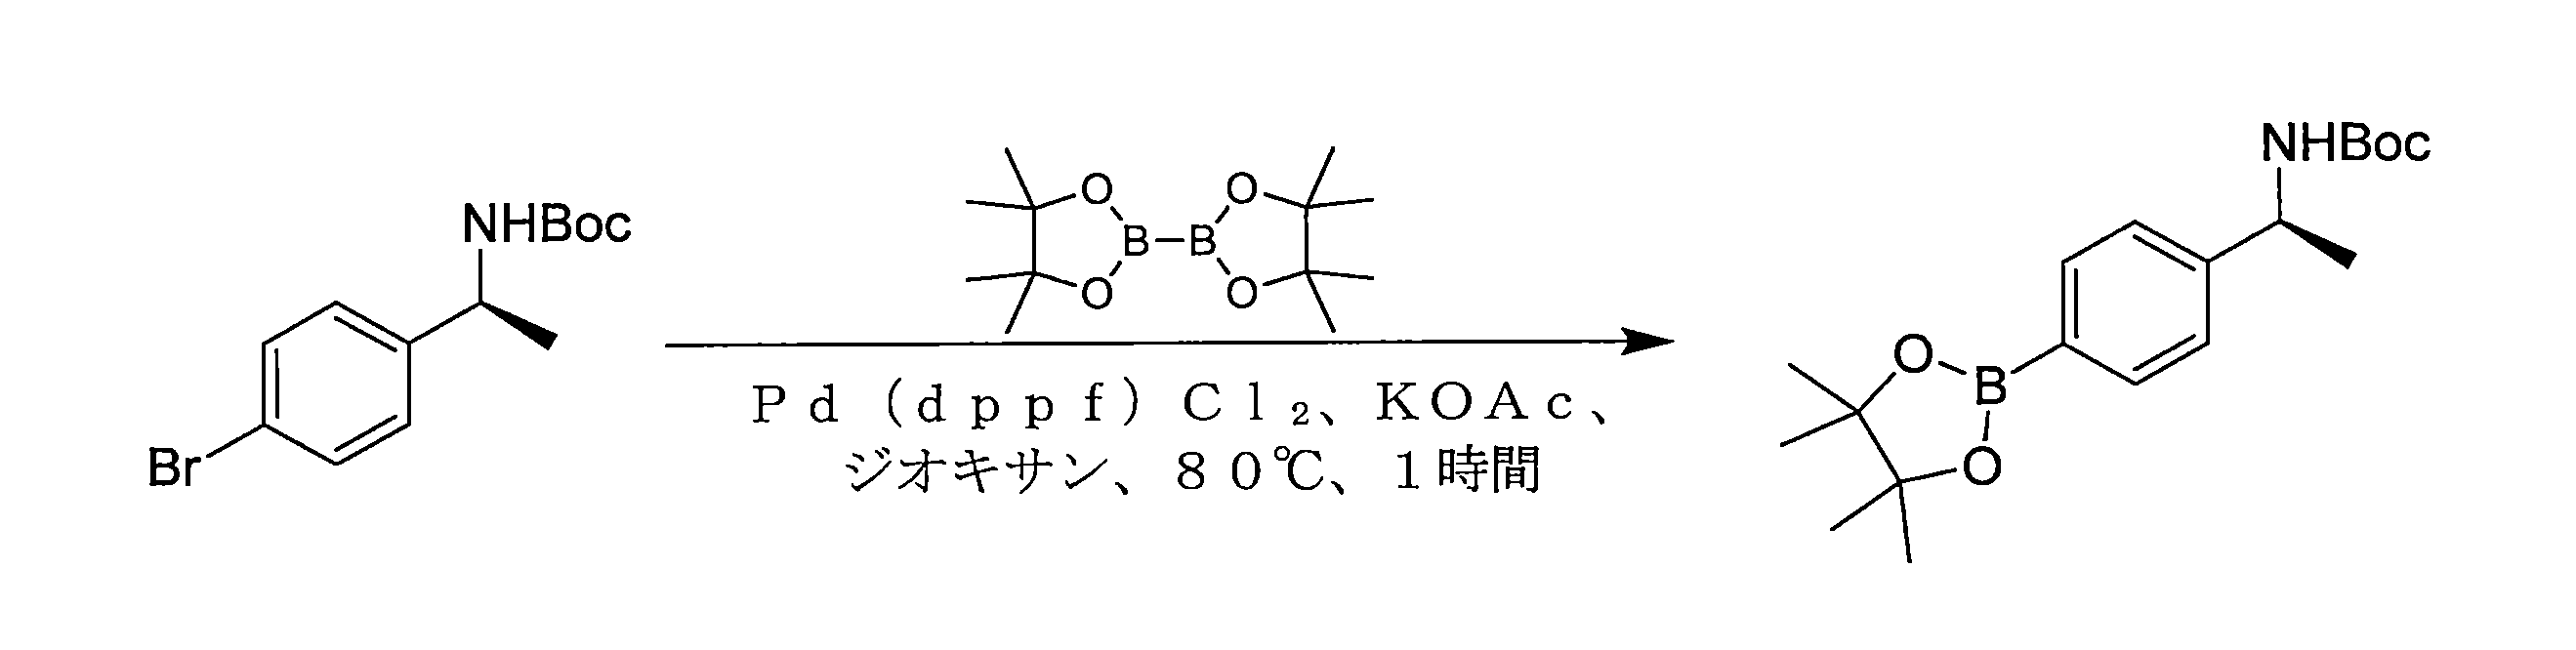
フラスコに、tert-ブチルN-[(1S)-1-(4-ブロモフェニル)エチル]カルバメート(1.4g、4.66mmol、1当量)、4,4,5,5-テトラメチル-2-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)-1,3,2-ジオキサボロラン(1.42g、5.60mmol、1.2当量)、(1,1’-ビス(ジフェニルホスフィノ)フェロセン)パラジウム(II)二塩化物(170mg、0.23mmol、0.05当量)、酢酸カリウム(915mg、9.33mmol、2当量)、およびジオキサン(30mL)を充填した。混合物を窒素で10分間パージし、次いで、80℃に1時間加熱した。反応混合物を20℃まで冷却し、セライトのパッドを通して濾過した。濾液を真空中で濃縮した。粗生成物をフラッシュクロマトグラフィー(石油エーテル:酢酸エチル=10:1)によって精製した。Tert-ブチルN-[(1S)-1-[4-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)フェニル]エチル]カルバメート(1.9g、粗製)を無色の油として得た。 A flask was charged with tert-butyl N-[(1S)-1-(4-bromophenyl)ethyl]carbamate (1.4 g, 4.66 mmol, 1 equiv.), 4,4,5,5-tetramethyl-2-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)-1,3,2-dioxaborolane (1.42 g, 5.60 mmol, 1.2 equiv.), (1,1'-bis(diphenylphosphino)ferrocene)palladium(II) dichloride (170 mg, 0.23 mmol, 0.05 equiv.), potassium acetate (915 mg, 9.33 mmol, 2 equiv.), and dioxane (30 mL). The mixture was purged with nitrogen for 10 min and then heated to 80 °C for 1 h. The reaction mixture was cooled to 20 °C and filtered through a pad of Celite. The filtrate was concentrated in vacuo. The crude product was purified by flash chromatography (petroleum ether:ethyl acetate=10:1). Tert-butyl N-[(1S)-1-[4-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)phenyl]ethyl]carbamate (1.9 g, crude) was obtained as a colorless oil.
工程4

ジオキサン(30mL)および水(5mL)中の5-ブロモ-4-(メトキシメチル)チアゾール(900mg、4.33mmol、1当量)およびtert-ブチルN-[(1S)-1-[4-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)フェニル]エチル]カルバメート(2.25g、6.49mmol、1.5当量)の混合物に、炭酸カリウム(1.20g、8.65mmol、2当量)および[1,1’-ビス(ジフェニルホスフィノ)フェロセン]ジクロロパラジウム(II)(316mg、0.43mmol、0.1当量)を窒素下で、15℃で一度に添加した。混合物を90℃で3時間撹拌した。混合物を15℃に冷却し、濾過し、真空下で濃縮した。残渣を、シリカゲルクロマトグラフィー(石油エーテル/酢酸エチル=10/1~1/1)によって精製し、tert-ブチルN-[(1S)-1-[4-[4-(メトキシメチル)チアゾール-5-イル]フェニル]エチル]カルバメート(1.87g、粗製)を黄色の固体として得た。 To a mixture of 5-bromo-4-(methoxymethyl)thiazole (900 mg, 4.33 mmol, 1 equiv.) and tert-butyl N-[(1S)-1-[4-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)phenyl]ethyl]carbamate (2.25 g, 6.49 mmol, 1.5 equiv.) in dioxane (30 mL) and water (5 mL), potassium carbonate (1.20 g, 8.65 mmol, 2 equiv.) and [1,1'-bis(diphenylphosphino)ferrocene]dichloropalladium(II) (316 mg, 0.43 mmol, 0.1 equiv.) were added in one portion at 15°C under nitrogen. The mixture was stirred at 90°C for 3 h. The mixture was cooled to 15°C, filtered, and concentrated under vacuum. The residue was purified by silica gel chromatography (petroleum ether/ethyl acetate = 10/1 to 1/1) to obtain tert-butyl N-[(1S)-1-[4-[4-(methoxymethyl)thiazol-5-yl]phenyl]ethyl]carbamate (1.87 g, crude) as a yellow solid.
例示的な化合物76について記載される手順を使用して、tert-ブチルN-[(1S)-1-[4-[4-(メトキシメチル)チアゾール-5-イル]フェニル]エチル]カルバメートtert-ブチルN-[(1S)-1-[4-[4-(ヒドロキシメチル)チアゾール-5-イル]フェニル]エチル]カルバメートを最終生成物に変換した。 Using the procedure described for exemplary compound 76, tert-butyl N-[(1S)-1-[4-[4-(methoxymethyl)thiazol-5-yl]phenyl]ethyl]carbamate tert-butyl N-[(1S)-1-[4-[4-(hydroxymethyl)thiazol-5-yl]phenyl]ethyl]carbamate was converted to the final product.
例示的な化合物184および185も、上に記載されるものと類似の手順を使用して調製した。 Exemplary compounds 184 and 185 were also prepared using procedures similar to those described above.
例示的な化合物170および134の例示的な合成

工程1

ジクロロメタン(5mL)中のメチル2-[3-(3-ヒドロキシプロピル)イソキサゾール-5-イル]-3-メチル-ブタノエート(460mg、1.91mmol、1当量)の溶液に、デスマーチンペルヨージナン(970mg、2.29mmol、1.2当量)を添加した。反応混合物を20℃で1時間撹拌した。酢酸エチル(50mL)を混合物に添加し、混合物を濾過し、濾液を真空中で濃縮した。有機相を水(20mL)およびブライン(20mL)洗浄し、硫酸ナトリウムで乾燥させ、濾過し、真空中で濃縮した。残渣を、シリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=100:1~50:1)によって精製した。メチル3-メチル-2-[3-(3-オキソプロピル)イソキサゾール-5-イル]ブタノエート(300mg、1.25mmol、収率65%)を無色の油として得た。 To a solution of methyl 2-[3-(3-hydroxypropyl)isoxazol-5-yl]-3-methyl-butanoate (460 mg, 1.91 mmol, 1 equiv.) in dichloromethane (5 mL) was added Dess-Martin periodinane (970 mg, 2.29 mmol, 1.2 equiv.). The reaction mixture was stirred at 20 °C for 1 h. Ethyl acetate (50 mL) was added to the mixture, the mixture was filtered, and the filtrate was concentrated in vacuum. The organic phase was washed with water (20 mL) and brine (20 mL), dried over sodium sulfate, filtered, and concentrated in vacuum. The residue was purified by silica gel chromatography (petroleum ether: ethyl acetate = 100: 1 to 50: 1). Methyl 3-methyl-2-[3-(3-oxopropyl)isoxazol-5-yl]butanoate (300 mg, 1.25 mmol, 65% yield) was obtained as a colorless oil.
他の実施例について上に記載される手順を使用して、以下のスキームに従って、メチル3-メチル-2-[3-(3-オキソプロピル)イソキサゾール-5-イル]ブタノエートを最終化合物に変換した。

例示的な化合物190および191を、上に記載されるものと類似の手順を使用して調製した。 Exemplary compounds 190 and 191 were prepared using procedures similar to those described above.
例示的な化合物112の例示的な合成

工程1

ジクロロメタン(150mL)中のブタ-3-イン-1-オール(5g、71.34mmol、1当量)の溶液に、p-トルエンスルホン酸ピリジニウム(1.9g、7.56mmol、0.1当量)および3,4-ジヒドロ-2H-ピラン(9.48g、112.70mmol、10.3mL、1.58当量)を15℃で添加し、混合物を15℃で18時間撹拌した。混合物を濃縮して、残渣を得た。残渣を、シリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=1:0~10:1)によって精製した。2-ブタ-3-イノキシテトラヒドロピラン(10.6g、68.74mmol、収率96%)を無色の油として得た。 To a solution of but-3-yn-1-ol (5 g, 71.34 mmol, 1 equiv.) in dichloromethane (150 mL), pyridinium p-toluenesulfonate (1.9 g, 7.56 mmol, 0.1 equiv.) and 3,4-dihydro-2H-pyran (9.48 g, 112.70 mmol, 10.3 mL, 1.58 equiv.) were added at 15°C, and the mixture was stirred at 15°C for 18 h. The mixture was concentrated to give a residue. The residue was purified by silica gel chromatography (petroleum ether:ethyl acetate=1:0-10:1). 2-but-3-ynoxytetrahydropyran (10.6 g, 68.74 mmol, 96% yield) was obtained as a colorless oil.
工程2

テトラヒドロフラン(20mL)中の2-ブタ-3-イノキシテトラヒドロピラン(2g、12.97mmol、1当量)の溶液に、n-ブチルリチウム(2.5M、5.7mL、1.1当量)を-78℃で添加し、混合物を-78℃で1時間撹拌した。次いで、パラホルムアルデヒド(2.10g、23.35mmol、1.8当量)を混合物に添加し、-78℃で30分間および20℃で15時間撹拌した。混合物を飽和塩化アンモニウム溶液(80mL)を用いてクエンチし、酢酸エチル(2×40mL)で抽出した。合わせた有機相を硫酸ナトリウムで乾燥させ、次いで、濃縮して、残渣を得た。残渣を、カラムクロマトグラフィー(石油エーテル:酢酸エチル=3:1~0:1)によって精製した。5-テトラヒドロピラン-2-イルオキシペント-2-イン-1-オール(1.4g、7.60mmol、収率58%)を無色の油として得た。 To a solution of 2-but-3-ynoxytetrahydropyran (2 g, 12.97 mmol, 1 eq.) in tetrahydrofuran (20 mL) was added n-butyllithium (2.5 M, 5.7 mL, 1.1 eq.) at −78°C, and the mixture was stirred at −78°C for 1 h. Then, paraformaldehyde (2.10 g, 23.35 mmol, 1.8 eq.) was added to the mixture and stirred at −78°C for 30 min and at 20°C for 15 h. The mixture was quenched with saturated ammonium chloride solution (80 mL) and extracted with ethyl acetate (2×40 mL). The combined organic phase was dried over sodium sulfate and then concentrated to give a residue. The residue was purified by column chromatography (petroleum ether:ethyl acetate=3:1 to 0:1). 5-Tetrahydropyran-2-yloxypent-2-yn-1-ol (1.4 g, 7.60 mmol, 58% yield) was obtained as a colorless oil.
工程3

テトラヒドロフラン(15mL)中の5-テトラヒドロピラン-2-イルオキシペント-2-イン-1-オール(1.4g、7.60mmol、1当量)およびp-トルエンスルホニルクロリド(2.90g、15.20mmol、2当量)の混合物に、水酸化カリウム(2.13g、38.00mmol、5当量)を添加した。次いで、反応混合物を20℃で2時間撹拌した。反応混合物を水(30mL)で希釈し、次いで、酢酸エチル(30mL×3)で抽出した。合わせた有機層をブライン(20mL×2)で洗浄し、硫酸ナトリウムで乾燥させ、濾過し、減圧下で濃縮して、残渣を得た。残渣を、カラムクロマトグラフィー(石油エーテル:酢酸エチル=50:1~10:1)によって精製した。化合物5-テトラヒドロピラン-2-イルオキペント-2-イニル4-メチルベンゼンスルホネート(2.5g、7.39mmol、収率97%)を無色の油として得た。 To a mixture of 5-tetrahydropyran-2-yloxypent-2-yn-1-ol (1.4 g, 7.60 mmol, 1 eq.) and p-toluenesulfonyl chloride (2.90 g, 15.20 mmol, 2 eq.) in tetrahydrofuran (15 mL), potassium hydroxide (2.13 g, 38.00 mmol, 5 eq.) was added. The reaction mixture was then stirred at 20°C for 2 h. The reaction mixture was diluted with water (30 mL) and then extracted with ethyl acetate (30 mL x 3). The combined organic layers were washed with brine (20 mL x 2), dried over sodium sulfate, filtered, and concentrated under reduced pressure to obtain a residue. The residue was purified by column chromatography (petroleum ether: ethyl acetate = 50: 1 to 10: 1). Compound 5-tetrahydropyran-2-yloxypent-2-ynyl 4-methylbenzenesulfonate (2.5 g, 7.39 mmol, 97% yield) was obtained as a colorless oil.
工程4

アセトニトリル(20mL)中の5-テトラヒドロピラン-2-イルオキシペント-2-イニル4-メチルベンゼンスルホネート(2.5g、7.39mmol、1当量)、4-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)-1H-ピラゾール(1.72g、8.86mmol、1.2当量)の溶液に、炭酸カリウム(2.04g、14.77mmol、2当量)を添加した。次いで、反応混合物を80℃で12時間撹拌した。反応混合物を水(30mL)で希釈し、次いで、酢酸エチル(30mL×3)で抽出した。合わせた有機層をブライン(20mL×2)で洗浄し、硫酸ナトリウムで乾燥させ、濾過し、減圧下で濃縮して、残渣を得た。残渣を、カラムクロマトグラフィー(石油エーテル:酢酸エチル=50:1~10:1)によって精製した。化合物1-(5-テトラヒドロピラン-2-イルオキシペント-2-イニル)-4-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)ピラゾール(2g、5.55mmol、収率75%)を淡い黄色の油として得た。 To a solution of 5-tetrahydropyran-2-yloxypent-2-ynyl 4-methylbenzenesulfonate (2.5 g, 7.39 mmol, 1 eq.), 4-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)-1H-pyrazole (1.72 g, 8.86 mmol, 1.2 eq.) in acetonitrile (20 mL), potassium carbonate (2.04 g, 14.77 mmol, 2 eq.) was added. The reaction mixture was then stirred at 80° C. for 12 h. The reaction mixture was diluted with water (30 mL) and then extracted with ethyl acetate (30 mL×3). The combined organic layers were washed with brine (20 mL×2), dried over sodium sulfate, filtered, and concentrated under reduced pressure to give a residue. The residue was purified by column chromatography (petroleum ether:ethyl acetate=50:1 to 10:1). The compound 1-(5-tetrahydropyran-2-yloxypent-2-ynyl)-4-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)pyrazole (2 g, 5.55 mmol, 75% yield) was obtained as a pale yellow oil.
他の実施例について上に記載される手順を使用して、以下のスキームに従って、1-(5-テトラヒドロピラン-2-イルオキシペント-2-イニル)-4-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)ピラゾールを最終化合物に変換した。

例示的な化合物131の例示的な合成

上記の他の実施例について記載され、当業者に明らかである手順を使用して、以下のスキームに従って、(2S,4R)-N-((R)-2-(ジメチルアミノ)-1-(4-(4-メチルチアゾール-5-イル)フェニル)-2-オキソエチル)-4-ヒドロキシピロリジン-2-カルボキサミドを調製した。

上記の他の実施例について記載される手順を使用して、(2S,4R)-N-((R)-2-(ジメチルアミノ)-1-(4-(4-メチルチアゾール-5-イル)フェニル)-2-オキソエチル)-4-ヒドロキシピロリジン-2-カルボキサミドを最終化合物に変換した。 Using the procedures described for other examples above, (2S,4R)-N-((R)-2-(dimethylamino)-1-(4-(4-methylthiazol-5-yl)phenyl)-2-oxoethyl)-4-hydroxypyrrolidine-2-carboxamide was converted to the final compound.
例示的な化合物171を、類似の手順を使用して調製した。 Exemplary compound 171 was prepared using a similar procedure.
例示的な化合物172の例示的な合成
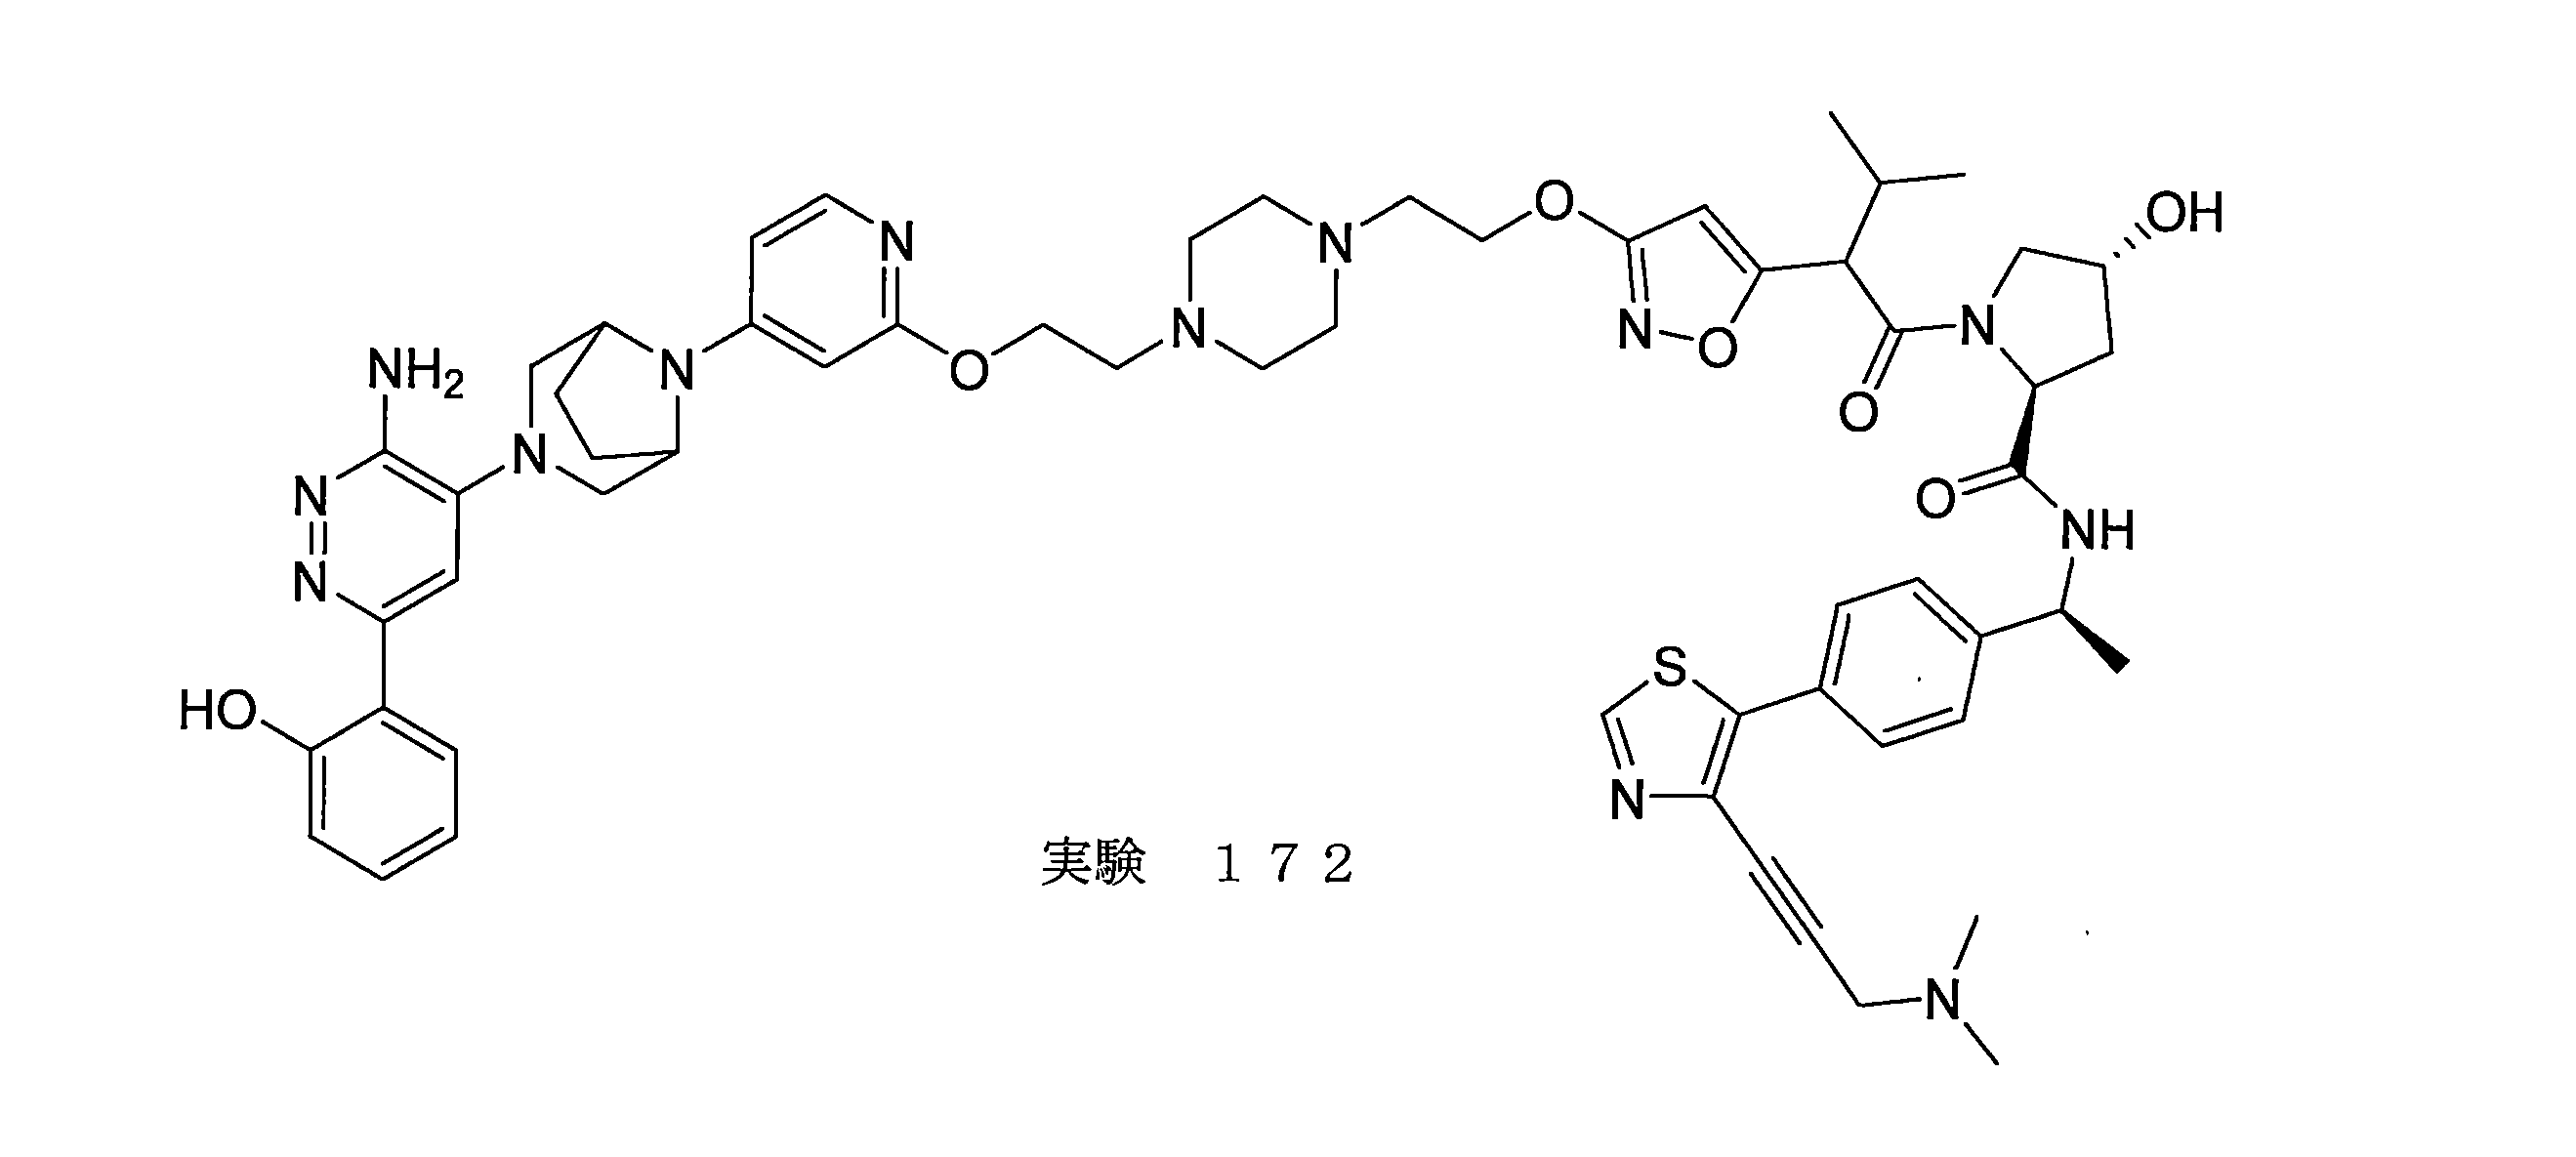
工程1
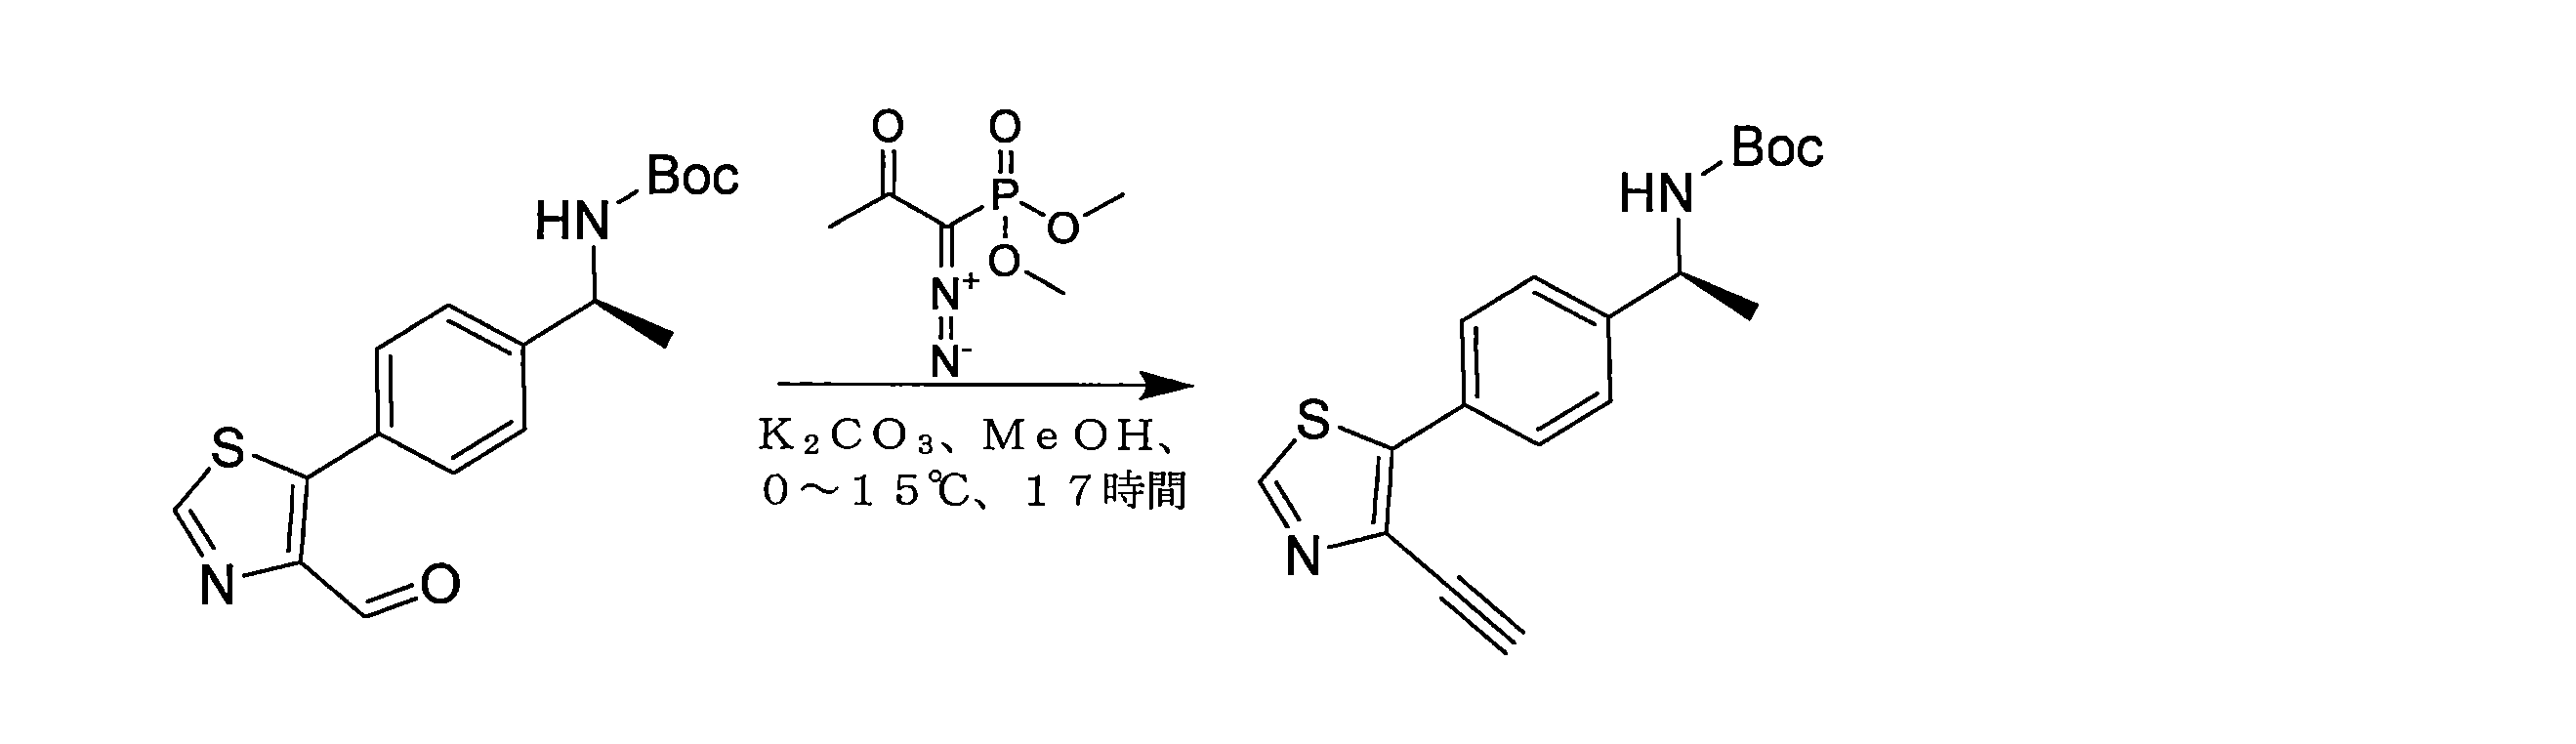
メタノール(20mL)中のtert-ブチルN-[(1S)-1-[4-(4-ホルミルチアゾール-5-イル)フェニル]エチル]カルバメート(2g、6.02mmol、1当量)および1-ジアゾ-1-ジメトキシホスホリル-プロパン-2-オン(1.16g、6.02mmol、1当量)の溶液に、炭酸カリウム(1.66g、12.03mmol、2当量)を0℃で添加した。反応混合物を15℃で16時間撹拌した。次いで、1-ジアゾ-1-ジメトキシホスホリル-プロパン-2-オン(116mg、0.60mmol、0.1当量)を添加し、反応混合物を15℃でさらに1時間撹拌した。酢酸エチル(50mL)および水(40mL)を添加し、混合物を分離した。有機層を硫酸ナトリウムで乾燥させ、次いで、真空下で濃縮して、残渣を得、これをシリカゲルカラムクロマトグラフィー(石油エーテル中10~30%酢酸エチル)によって精製して、tert-ブチルN-[(1S)-1-[4-(4-エチニルチアゾール-5-イル)フェニル]エチル]カルバメート(1.8g、5.48mmol、収率91%)を白色の固体として得た。 To a solution of tert-butyl N-[(1S)-1-[4-(4-formylthiazol-5-yl)phenyl]ethyl]carbamate (2 g, 6.02 mmol, 1 eq.) and 1-diazo-1-dimethoxyphosphoryl-propan-2-one (1.16 g, 6.02 mmol, 1 eq.) in methanol (20 mL) was added potassium carbonate (1.66 g, 12.03 mmol, 2 eq.) at 0 °C. The reaction mixture was stirred at 15 °C for 16 h. Then 1-diazo-1-dimethoxyphosphoryl-propan-2-one (116 mg, 0.60 mmol, 0.1 eq.) was added and the reaction mixture was stirred at 15 °C for an additional 1 h. Ethyl acetate (50 mL) and water (40 mL) were added and the mixture was separated. The organic layer was dried over sodium sulfate and then concentrated under vacuum to give a residue that was purified by silica gel column chromatography (10-30% ethyl acetate in petroleum ether) to give tert-butyl N-[(1S)-1-[4-(4-ethynylthiazol-5-yl)phenyl]ethyl]carbamate (1.8 g, 5.48 mmol, 91% yield) as a white solid.
工程2

ジオキサン(8mL)中のtert-ブチルN-[(1S)-1-[4-(4-エチニルチアゾール-5-イル)フェニル]エチル]カルバメート(400mg、1.22mmol、1当量)、ジメチルアミン(1.37g、12.18mmol、1.54mL、10当量)、およびトリホルモール(362mg、4.02mmol、3.3当量)の溶液に、ヨウ化銅(232mg、1.22mmol、1当量)を0℃で添加した。反応混合物を脱気し、窒素を3回充填し、次いで、30℃で4時撹拌した。反応溶液を真空下で濃縮して、残渣を得た。残渣を、シリカゲルカラムクロマトグラフィー(ジクロロメタン中0~8%メタノール)によって精製して、tert-ブチルN-[(1S)-1-[4-[4-[3-(ジメチルアミノ)プロパ-1-イニル]チアゾール-5-イル]フェニル]エチル]カルバメート(274mg、0.71mmol、収率58%)を淡い黄色のガムとして得た。 To a solution of tert-butyl N-[(1S)-1-[4-(4-ethynylthiazol-5-yl)phenyl]ethyl]carbamate (400 mg, 1.22 mmol, 1 equiv.), dimethylamine (1.37 g, 12.18 mmol, 1.54 mL, 10 equiv.), and triformol (362 mg, 4.02 mmol, 3.3 equiv.) in dioxane (8 mL) was added copper iodide (232 mg, 1.22 mmol, 1 equiv.) at 0°C. The reaction mixture was degassed and filled with nitrogen three times, then stirred at 30°C for 4 h. The reaction solution was concentrated under vacuum to give a residue. The residue was purified by silica gel column chromatography (0-8% methanol in dichloromethane) to give tert-butyl N-[(1S)-1-[4-[4-[3-(dimethylamino)prop-1-ynyl]thiazol-5-yl]phenyl]ethyl]carbamate (274 mg, 0.71 mmol, 58% yield) as a pale yellow gum.
例示的な化合物76について記載される手順を使用して、tert-ブチルN-[(1S)-1-[4-[4-[3-(ジメチルアミノ)プロパ-1-イニル]チアゾール-5-イル]フェニル]エチル]カルバメートを最終化合物に変換した。 Using the procedure described for exemplary compound 76, tert-butyl N-[(1S)-1-[4-[4-[3-(dimethylamino)prop-1-ynyl]thiazol-5-yl]phenyl]ethyl]carbamate was converted to the final compound.
例示的な化合物132の例示的な合成

工程1

アセトニトリル(200mL)中の2,2-ジエトキシエタノール(22.87g、170mmol、1.5当量)および4-ブロモ-2-フルオロ-ピリジン(20g、113.65mmol、1当量)の混合物に、炭酸セシウム(74.06g、227mmol、2当量)を窒素下で、15℃で一度に添加した。混合物を90℃で3時間撹拌した。混合物を20℃に冷却し、濾過し、真空下で濃縮した。残渣を、シリカゲルクロマトグラフィー(石油エーテル)によって精製して、4-ブロモ-2-(2,2-ジエトキシエトキシ)ピリジン(22.8g、74.7mmol、収率66%)を黄色の油として得た。 To a mixture of 2,2-diethoxyethanol (22.87 g, 170 mmol, 1.5 equiv.) and 4-bromo-2-fluoro-pyridine (20 g, 113.65 mmol, 1 equiv.) in acetonitrile (200 mL), cesium carbonate (74.06 g, 227 mmol, 2 equiv.) was added in one portion under nitrogen at 15°C. The mixture was stirred at 90°C for 3 h. The mixture was cooled to 20°C, filtered, and concentrated under vacuum. The residue was purified by silica gel chromatography (petroleum ether) to give 4-bromo-2-(2,2-diethoxyethoxy)pyridine (22.8 g, 74.7 mmol, 66% yield) as a yellow oil.
工程2
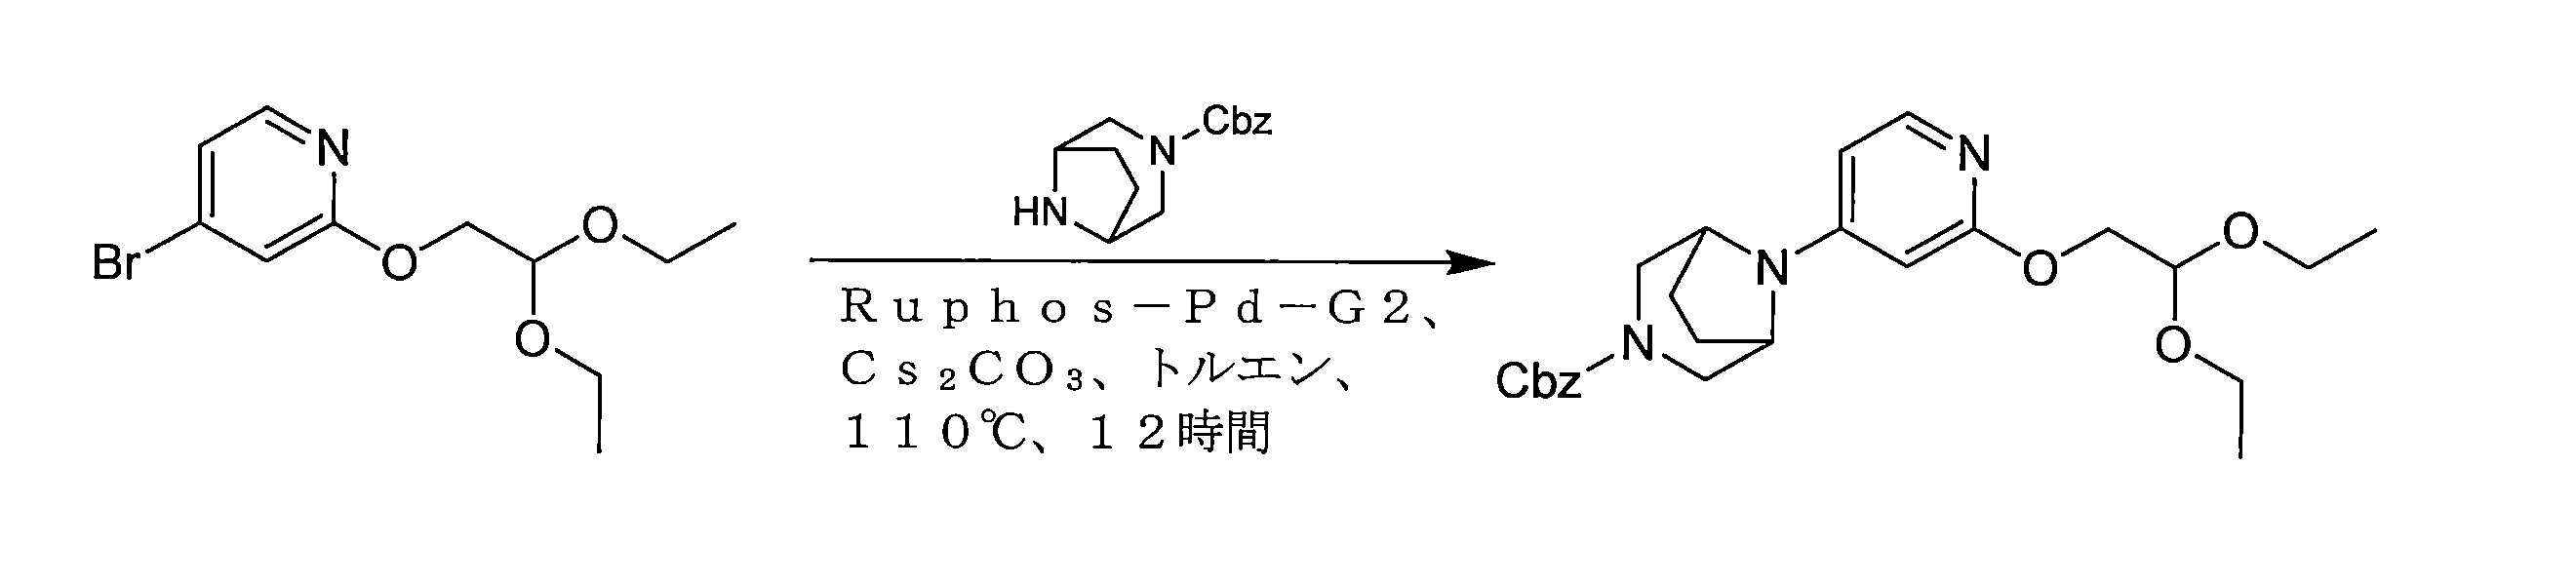
トルエン(200mL)中の4-ブロモ-2-(2,2-ジエトキシエトキシ)ピリジン(10g、34.46mmol、1当量)およびベンジル3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレート(8.49g、34.46mmol、1当量)の混合物に、炭酸セシウム(22.46g、68.93mmol、2当量)および[2-(2-アミノフェニル)フェニル]-クロロ-パラジウムジシクロヘキシル-[2-(2,6-ジイソプロポキシフェニル)フェニル]ホスファン(1.61g、2.07mmol、0.06当量)を窒素下で、15℃で一度に添加した。混合物を110℃で12時間撹拌した。混合物を20℃に冷却し、濾過し、真空下で濃縮した。残渣を、シリカゲルクロマトグラフィー(石油エーテル/酢酸エチル=10/1、3/1)によって精製して、ベンジル8-[2-(2,2-ジエトキシエトキシ)-4-ピリジル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレート(12.1g、26.6mmol、収率77%)を黄色の油として得た。 To a mixture of 4-bromo-2-(2,2-diethoxyethoxy)pyridine (10 g, 34.46 mmol, 1 eq.) and benzyl 3,8-diazabicyclo[3.2.1]octane-3-carboxylate (8.49 g, 34.46 mmol, 1 eq.) in toluene (200 mL), cesium carbonate (22.46 g, 68.93 mmol, 2 eq.) and [2-(2-aminophenyl)phenyl]-chloro-palladium dicyclohexyl-[2-(2,6-diisopropoxyphenyl)phenyl]phosphane (1.61 g, 2.07 mmol, 0.06 eq.) were added in one portion under nitrogen at 15 °C. The mixture was stirred at 110 °C for 12 h. The mixture was cooled to 20 °C, filtered, and concentrated under vacuum. The residue was purified by silica gel chromatography (petroleum ether/ethyl acetate = 10/1, 3/1) to obtain benzyl 8-[2-(2,2-diethoxyethoxy)-4-pyridyl]-3,8-diazabicyclo[3.2.1]octane-3-carboxylate (12.1 g, 26.6 mmol, 77% yield) as a yellow oil.
工程3
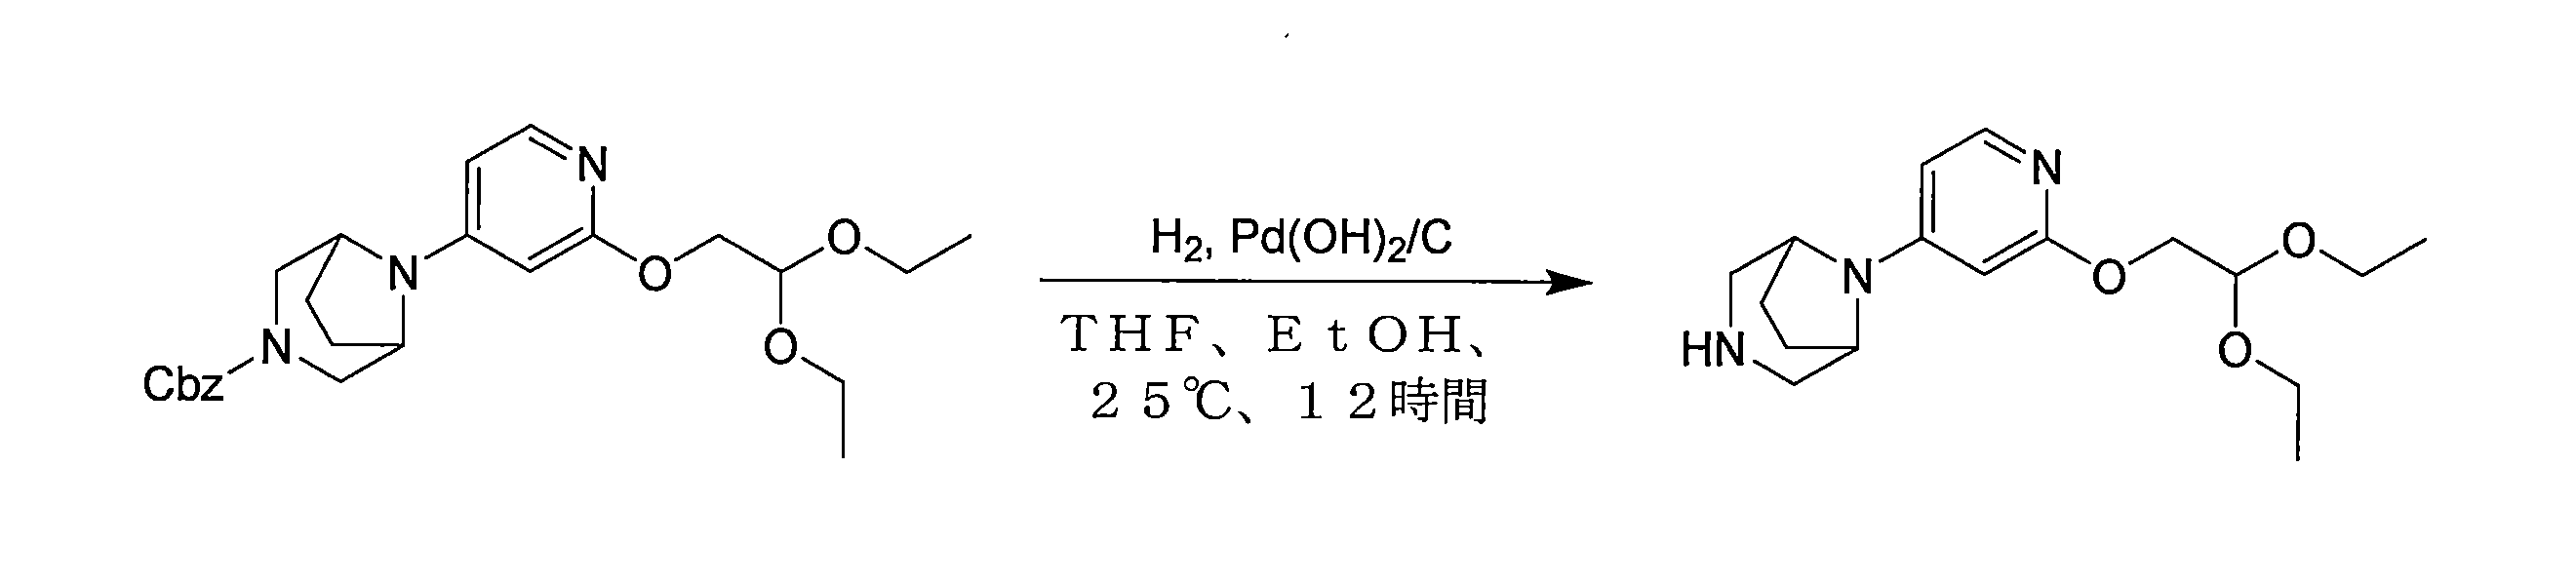
テトラヒドロフラン(300mL)およびエタノール(300mL)中のベンジル8-[2-(2,2-ジエトキシエトキシ)-4-ピリジル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレート(13.1g、28.76mmol、1当量)の溶液に、10%水酸化パラジウム炭素(4.04g、2.88mmol、0.1当量)を窒素下で添加した。懸濁液を真空下で脱気し、水素で数回パージした。混合物を、水素(50psi)下で、60℃で12時間撹拌した。反応混合物を濾過し、濾液を濃縮した。粗8-[2-(2,2-ジエトキシエトキシ)-4-ピリジル]-3,8-ジアザビシクロ[3.2.1]オクタン(8.27g、25.73mmol、収率89.47%)を茶色の油として得た。 To a solution of benzyl 8-[2-(2,2-diethoxyethoxy)-4-pyridyl]-3,8-diazabicyclo[3.2.1]octane-3-carboxylate (13.1 g, 28.76 mmol, 1 equiv.) in tetrahydrofuran (300 mL) and ethanol (300 mL) was added 10% palladium hydroxide on carbon (4.04 g, 2.88 mmol, 0.1 equiv.) under nitrogen. The suspension was degassed under vacuum and purged with hydrogen several times. The mixture was stirred under hydrogen (50 psi) at 60 °C for 12 h. The reaction mixture was filtered and the filtrate was concentrated. Crude 8-[2-(2,2-diethoxyethoxy)-4-pyridyl]-3,8-diazabicyclo[3.2.1]octane (8.27 g, 25.73 mmol, 89.47% yield) was obtained as a brown oil.
上記の他の実施例について記載される手順、ならびに当業者に明らかである標準的な手順を使用して、以下のスキームに従って、8-[2-(2,2-ジエトキシエトキシ)-4-ピリジル]-3,8-ジアザビシクロ[3.2.1]オクタンを最終化合物に変換した。

例示的な化合物132について記載されるものと類似の手順を使用して、以下の例示的な化合物を調製した:135、136、139、141、133、140、142、および210。 Using procedures similar to those described for exemplary compound 132, the following exemplary compounds were prepared: 135, 136, 139, 141, 133, 140, 142, and 210.
例示的な化合物173および102の例示的な合成
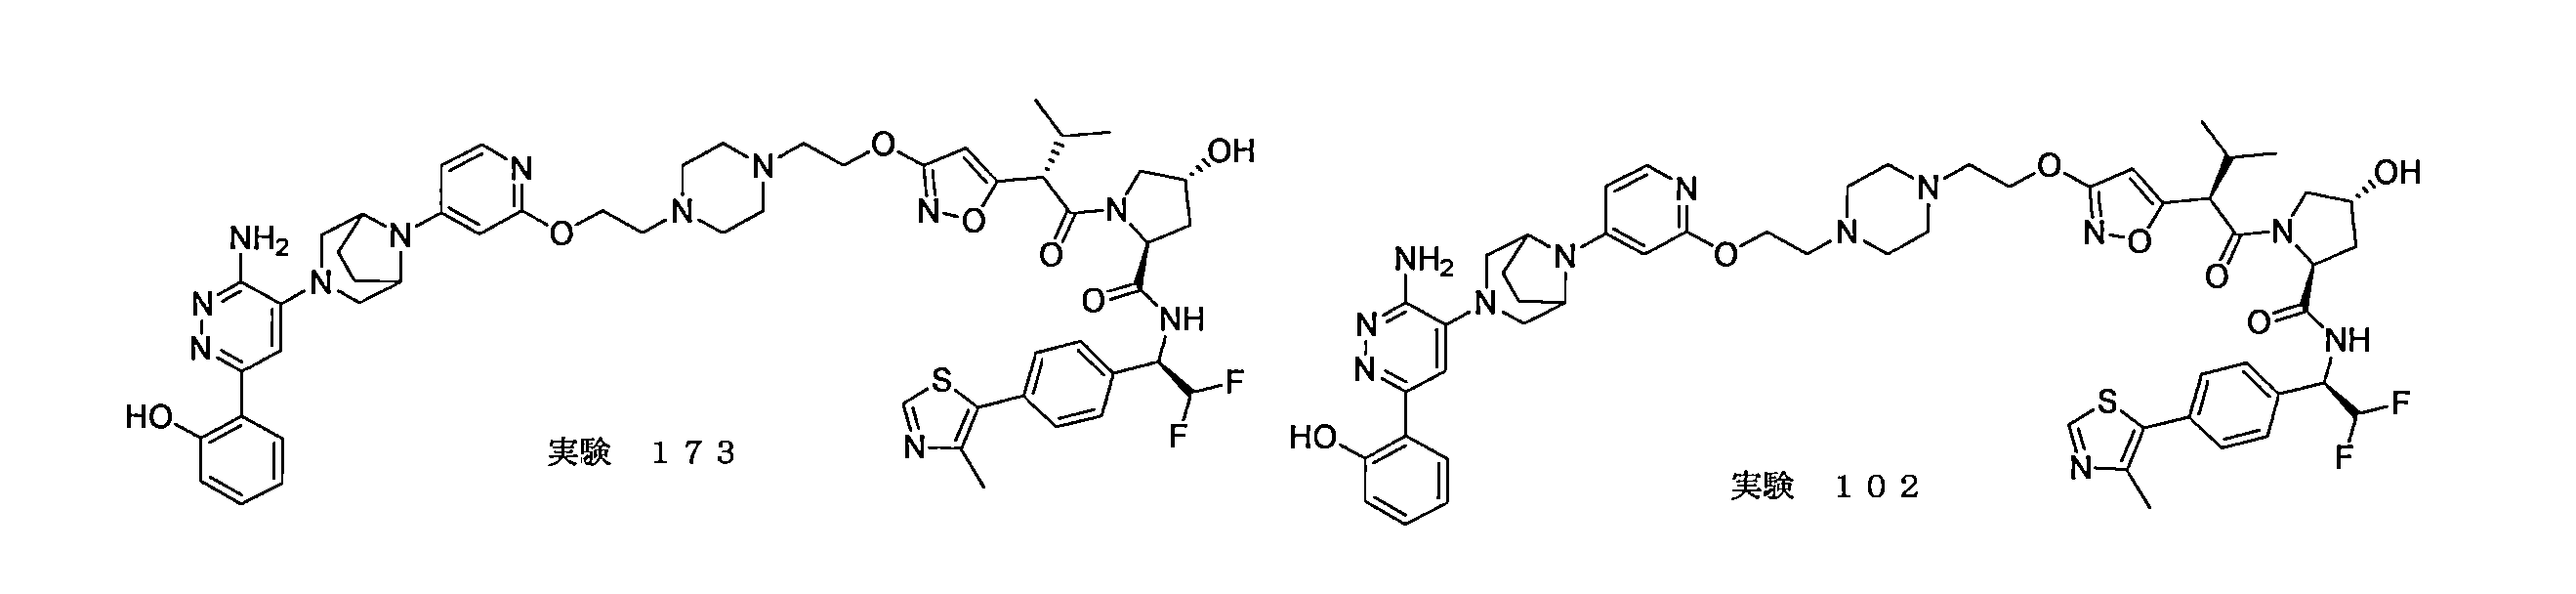
工程1

窒素不活性雰囲気下でパージし、維持した250mLの三口丸底フラスコに、THF(150.0mL)中の1,4-ジブロモベンゼン(12.0g、50.8mmol、1.0当量)を-78℃で入れた。次いで、n-ブチルリチウム溶液(ヘキサン中1M、60.9mL、60.9mmol、1.2当量)を-78℃で添加した。得られた混合物を-78℃で30分間撹拌し、次いで、2,2-ジフルオロ酢酸エチル(12.6g、101.5mmol、2.0当量)を-78℃でゆっくり添加した。冷却浴を除去し、得られた混合物を室温でさらに1時間撹拌した。次いで、反応物を、50mLの水を添加することによってクエンチした。得られた混合物を酢酸エチル(250mL×3)で抽出し、有機層を合わせ、ブラインで洗浄し、無水硫酸ナトリウムで乾燥させ、減圧下で濃縮した。残渣を、酢酸エチル/石油エーテル(1:10)で溶出するシリカゲルカラム上に適用した。これにより、7.6g(64%)の1-(4-ブロモフェニル)-2,2-ジフルオロエタン-1-オンを淡い黄色の半固体として得た。 A 250 mL three-neck round bottom flask purged and maintained under nitrogen inert atmosphere was charged with 1,4-dibromobenzene (12.0 g, 50.8 mmol, 1.0 equiv) in THF (150.0 mL) at -78 °C. Then, n-butyllithium solution (1 M in hexanes, 60.9 mL, 60.9 mmol, 1.2 equiv) was added at -78 °C. The resulting mixture was stirred at -78 °C for 30 minutes, and then 2,2-difluoroethyl acetate (12.6 g, 101.5 mmol, 2.0 equiv) was added slowly at -78 °C. The cooling bath was removed and the resulting mixture was stirred at room temperature for an additional hour. The reaction was then quenched by adding 50 mL of water. The resulting mixture was extracted with ethyl acetate (250 mL x 3), and the organic layers were combined, washed with brine, dried over anhydrous sodium sulfate, and concentrated under reduced pressure. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (1:10). This gave 7.6 g (64%) of 1-(4-bromophenyl)-2,2-difluoroethan-1-one as a pale yellow semi-solid.
工程2
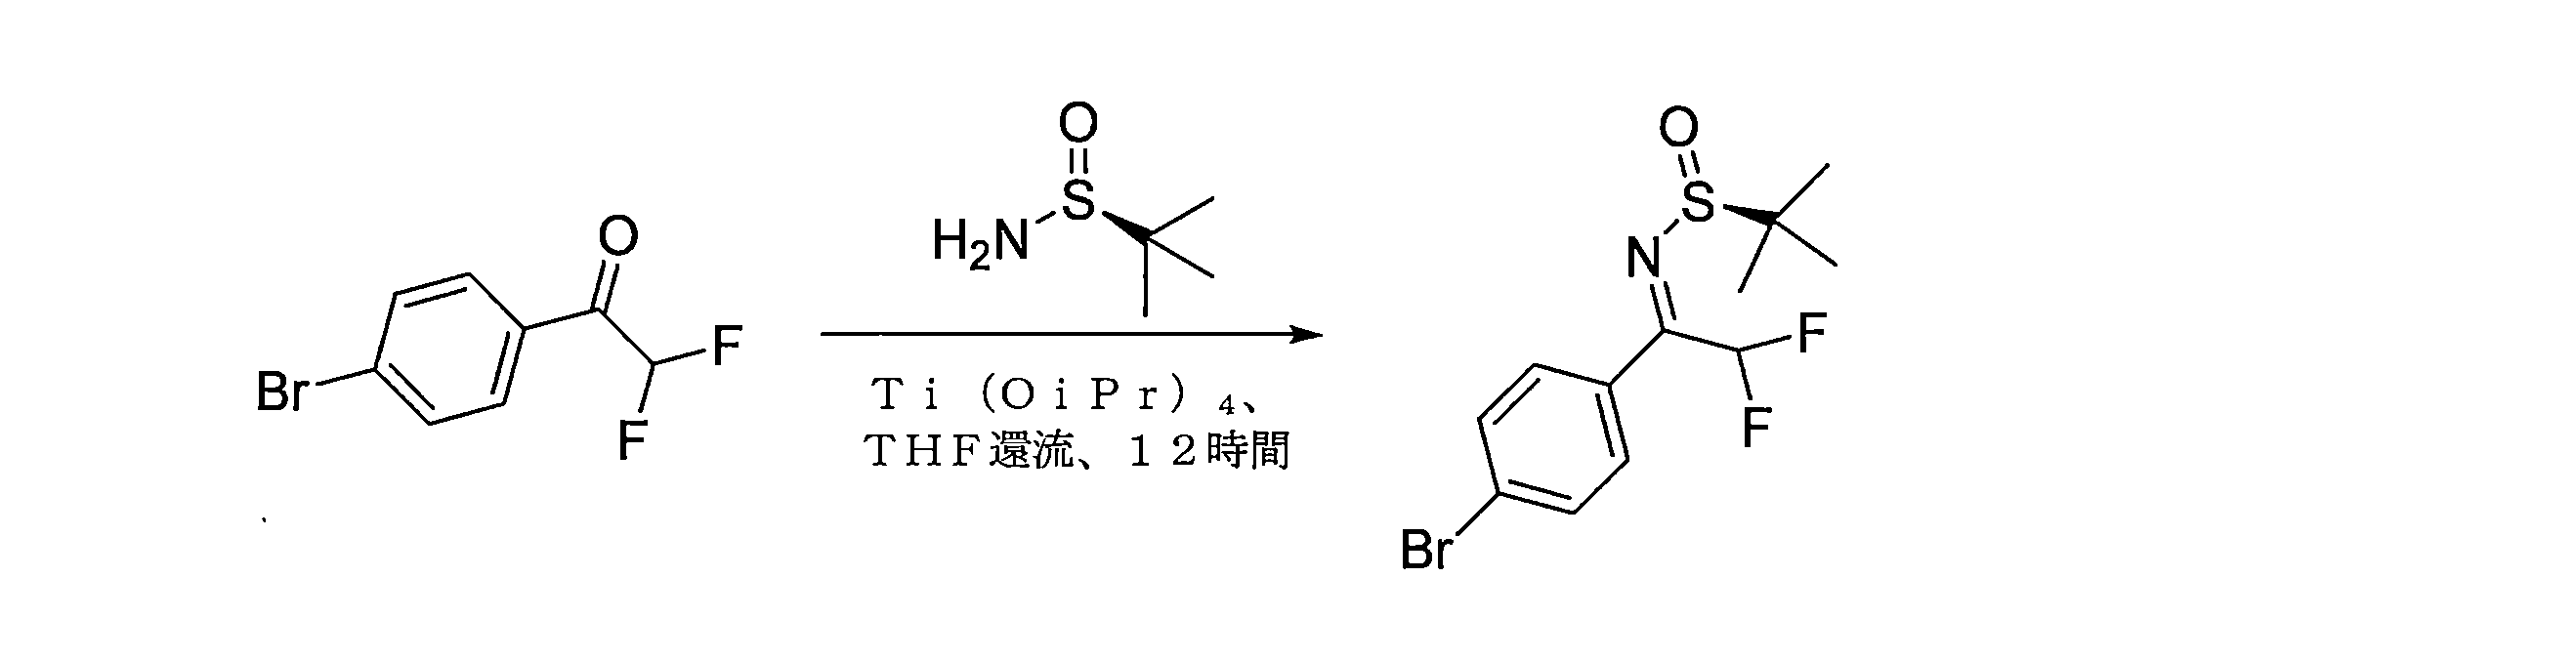
窒素の不活性雰囲気下でパージし、維持した30mLの密封チューブに、THF(10mL)中の1-(4-ブロモフェニル)-2,2-ジフルオロエタン-1-オン(3.5g、14.9mmol、1.0当量)、(S)-2-メチルプロパン-2-スルフィンアミド(3.6g、29.8mmol、2.0当量)、およびテトラキス(プロパン-2-イルオキシ)チタン(12.7g、44.7mmol、3.0当量)を窒素雰囲気下で入れた。得られた混合物を、窒素雰囲気下で、油浴中で、80℃で14時間撹拌した。混合物を減圧下で濃縮した。残渣を、酢酸エチル/石油エーテル(1:5)で溶出するシリカゲルカラム上に適用した。これにより、2.8g(56%)の(S)-N-[(1Z)-1-(4-ブロモフェニル)-2,2-ジフルオロエチリデン]-2-メチルプロパン-2-スルフィンアミドを黄色の油として得た。 In a 30 mL sealed tube purged and maintained under an inert atmosphere of nitrogen, 1-(4-bromophenyl)-2,2-difluoroethan-1-one (3.5 g, 14.9 mmol, 1.0 equiv.), (S)-2-methylpropane-2-sulfinamide (3.6 g, 29.8 mmol, 2.0 equiv.), and tetrakis(propan-2-yloxy)titanium (12.7 g, 44.7 mmol, 3.0 equiv.) in THF (10 mL) were placed under nitrogen atmosphere. The resulting mixture was stirred at 80 °C in an oil bath under nitrogen atmosphere for 14 h. The mixture was concentrated under reduced pressure. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (1:5). This yielded 2.8 g (56%) of (S)-N-[(1Z)-1-(4-bromophenyl)-2,2-difluoroethylidene]-2-methylpropane-2-sulfinamide as a yellow oil.
工程3
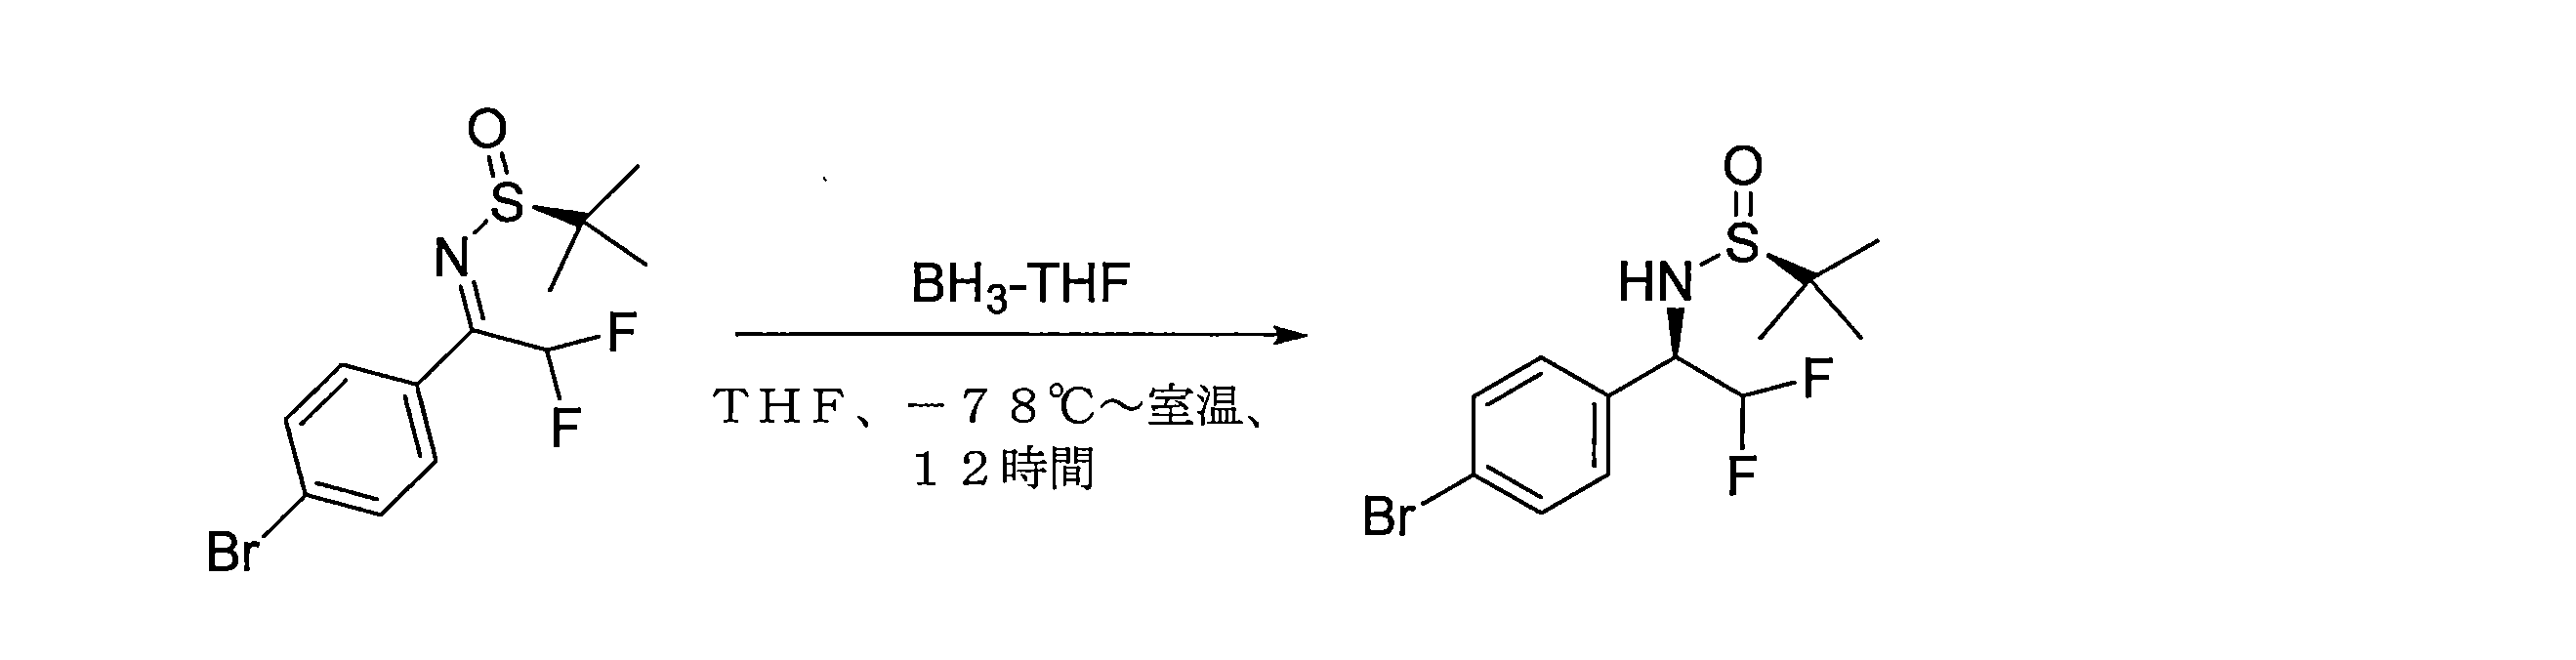
窒素不活性雰囲気下でパージし、維持した250mLの三口丸底フラスコに、THF(40.0mL)中の(S)-N-[(1Z)-1-(4-ブロモフェニル)-2,2-ジフルオロエチリデン]-2-メチルプロパン-2-スルフィンアミド(2.8g、8.3mmol、1当量)およびBH3-THF(40.0mL、40.0mmol、1.0M、4.8当量)の溶液を0℃で入れた。得られた混合物を0℃で1時間撹拌し、次いで、室温に温め、室温でさらに6時間撹拌した。反応を、100mLの水の添加により停止させた。得られた混合物を酢酸エチル(100mL×2)で抽出した。有機層を合わせ、減圧下で濃縮した。残渣を、ジクロロメタン/石油エーテル(1:1)で溶出するシリカゲルカラム上に適用した。これにより、1.3g(45%)の(S)-N-[(1R)-1-(4-ブロモフェニル)-2,2-ジフルオロエチル]-2-メチルプロパン-2-スルフィンアミドを白色の固体として得た。 A solution of (S)-N-[(1Z)-1-(4-bromophenyl)-2,2-difluoroethylidene]-2-methylpropane-2-sulfinamide (2.8 g, 8.3 mmol, 1 equiv.) and BH3-THF (40.0 mL, 40.0 mmol, 1.0 M, 4.8 equiv.) in THF (40.0 mL) was placed in a 250 mL three-neck round bottom flask purged and maintained under nitrogen inert atmosphere at 0°C. The resulting mixture was stirred at 0°C for 1 h, then warmed to room temperature and stirred at room temperature for an additional 6 h. The reaction was quenched by the addition of 100 mL of water. The resulting mixture was extracted with ethyl acetate (100 mL x 2). The organic layers were combined and concentrated under reduced pressure. The residue was applied onto a silica gel column eluting with dichloromethane/petroleum ether (1:1). This yielded 1.3 g (45%) of (S)-N-[(1R)-1-(4-bromophenyl)-2,2-difluoroethyl]-2-methylpropane-2-sulfinamide as a white solid.
工程4
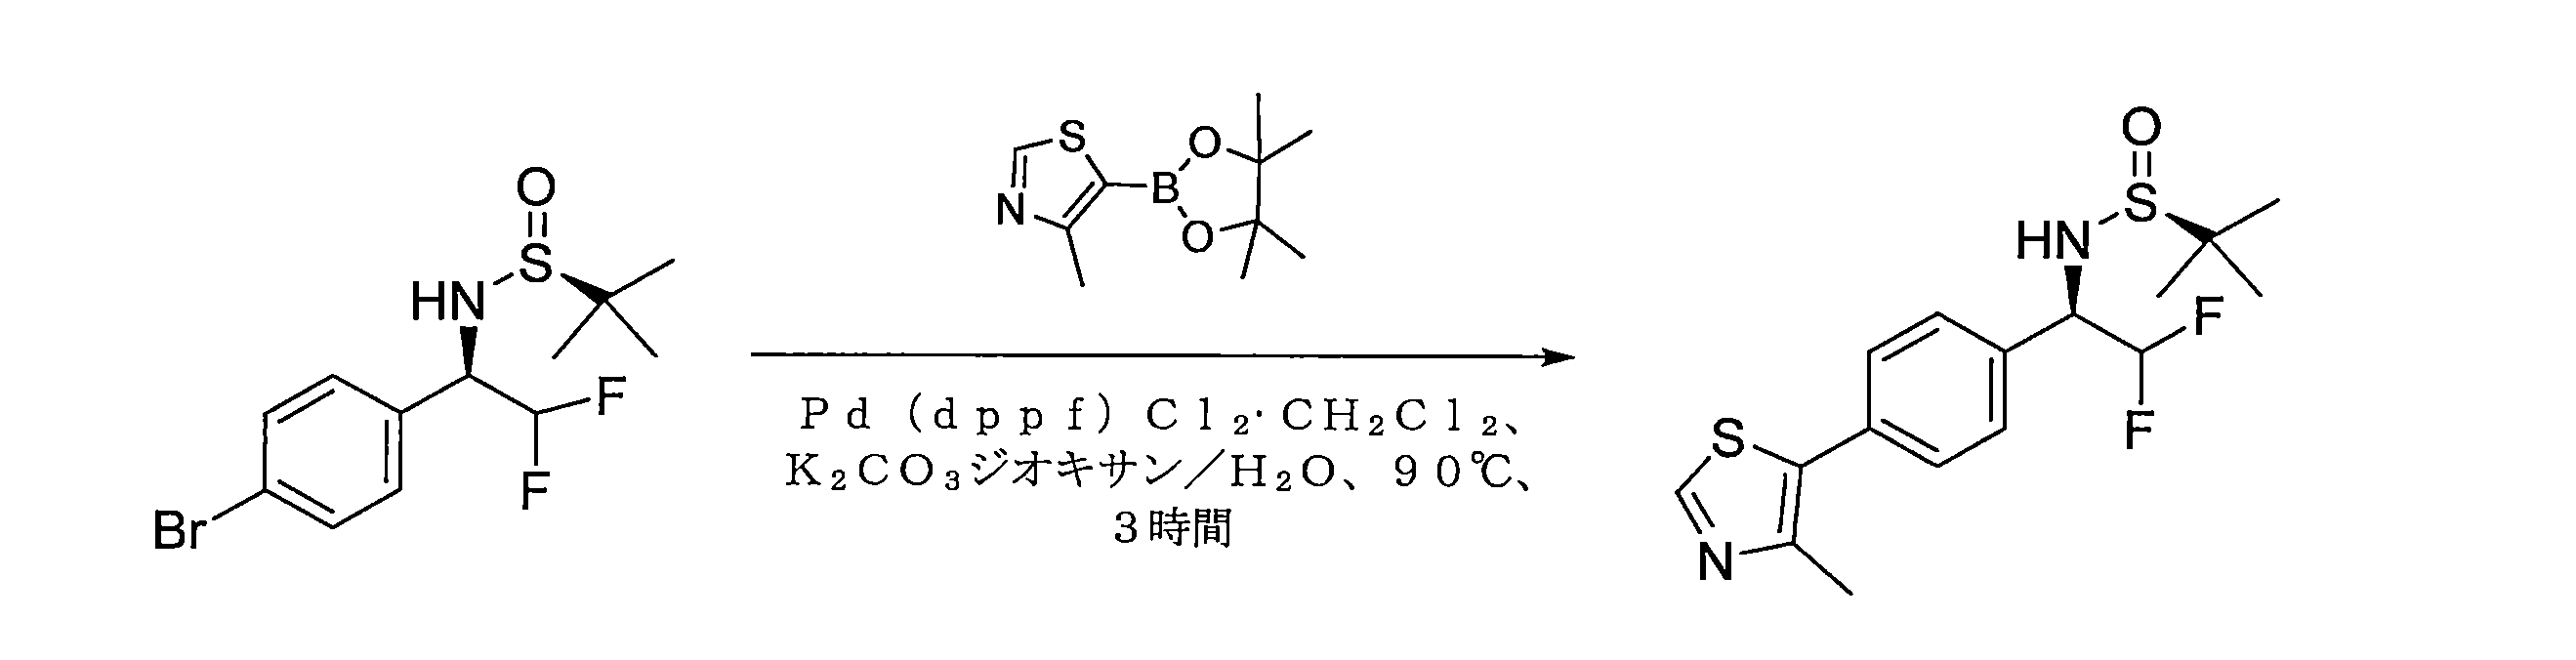
窒素の不活性雰囲気下でパージし、維持した30mLの密封チューブに、ジオキサン(15.0mL)および水(3.0mL)中の(S)-N-[(1R)-1-(4-ブロモフェニル)-2,2-ジフルオロエチル]-2-メチルプロパン-2-スルフィンアミド(1.3g、3.8mmol、1.0当量)、4-メチル-5-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)-1,3-チアゾール(1.25g、5.5mmol、1.5当量)、K2CO3(1.5g、11.3mmol、3.0当量)、およびPd(dppf)Cl2.CH2Cl2(62.0mg、0.08mmol、0.02当量)を窒素雰囲気下で入れた。得られた懸濁液を、窒素雰囲気下で、油浴中で、90℃で12時間撹拌した。混合物を減圧下で濃縮した。残渣を、酢酸エチル/石油エーテル(1:1)で溶出するシリカゲルカラム上に適用した。これにより、750.0mg(56%)の(S)-N-[(1R)-2,2-ジフルオロ-1-[4-(4-メチル-1,3-チアゾール-5-イル)-フェニル]エチル]-2-メチルプロパン-2-スルフィンアミドを淡い黄色の固体として得た。 A 30 mL sealed tube purged and maintained under an inert atmosphere of nitrogen was charged with (S)-N-[(1R)-1-(4-bromophenyl)-2,2-difluoroethyl]-2-methylpropane-2-sulfinamide (1.3 g, 3.8 mmol, 1.0 equiv.), 4-methyl-5-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)-1,3-thiazole (1.25 g, 5.5 mmol, 1.5 equiv.), K2CO3 (1.5 g, 11.3 mmol, 3.0 equiv.), and Pd(dppf)Cl2. CH2Cl2 (62.0 mg, 0.08 mmol, 0.02 equiv.) in dioxane (15.0 mL) and water (3.0 mL) under nitrogen atmosphere. The resulting suspension was stirred at 90 °C for 12 h in an oil bath under nitrogen atmosphere. The mixture was concentrated under reduced pressure. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (1:1). This gave 750.0 mg (56%) of (S)-N-[(1R)-2,2-difluoro-1-[4-(4-methyl-1,3-thiazol-5-yl)-phenyl]ethyl]-2-methylpropane-2-sulfinamide as a pale yellow solid.
この出願における他の実施例について記載されるものと類似の手順を使用して、以下のスキームに従って、(S)-N-[(1R)-2,2-ジフルオロ-1-[4-(4-メチル-1,3-チアゾール-5-イル)-フェニル]エチル]-2-メチルプロパン-2-スルフィンアミドを最終化合物に変換した。
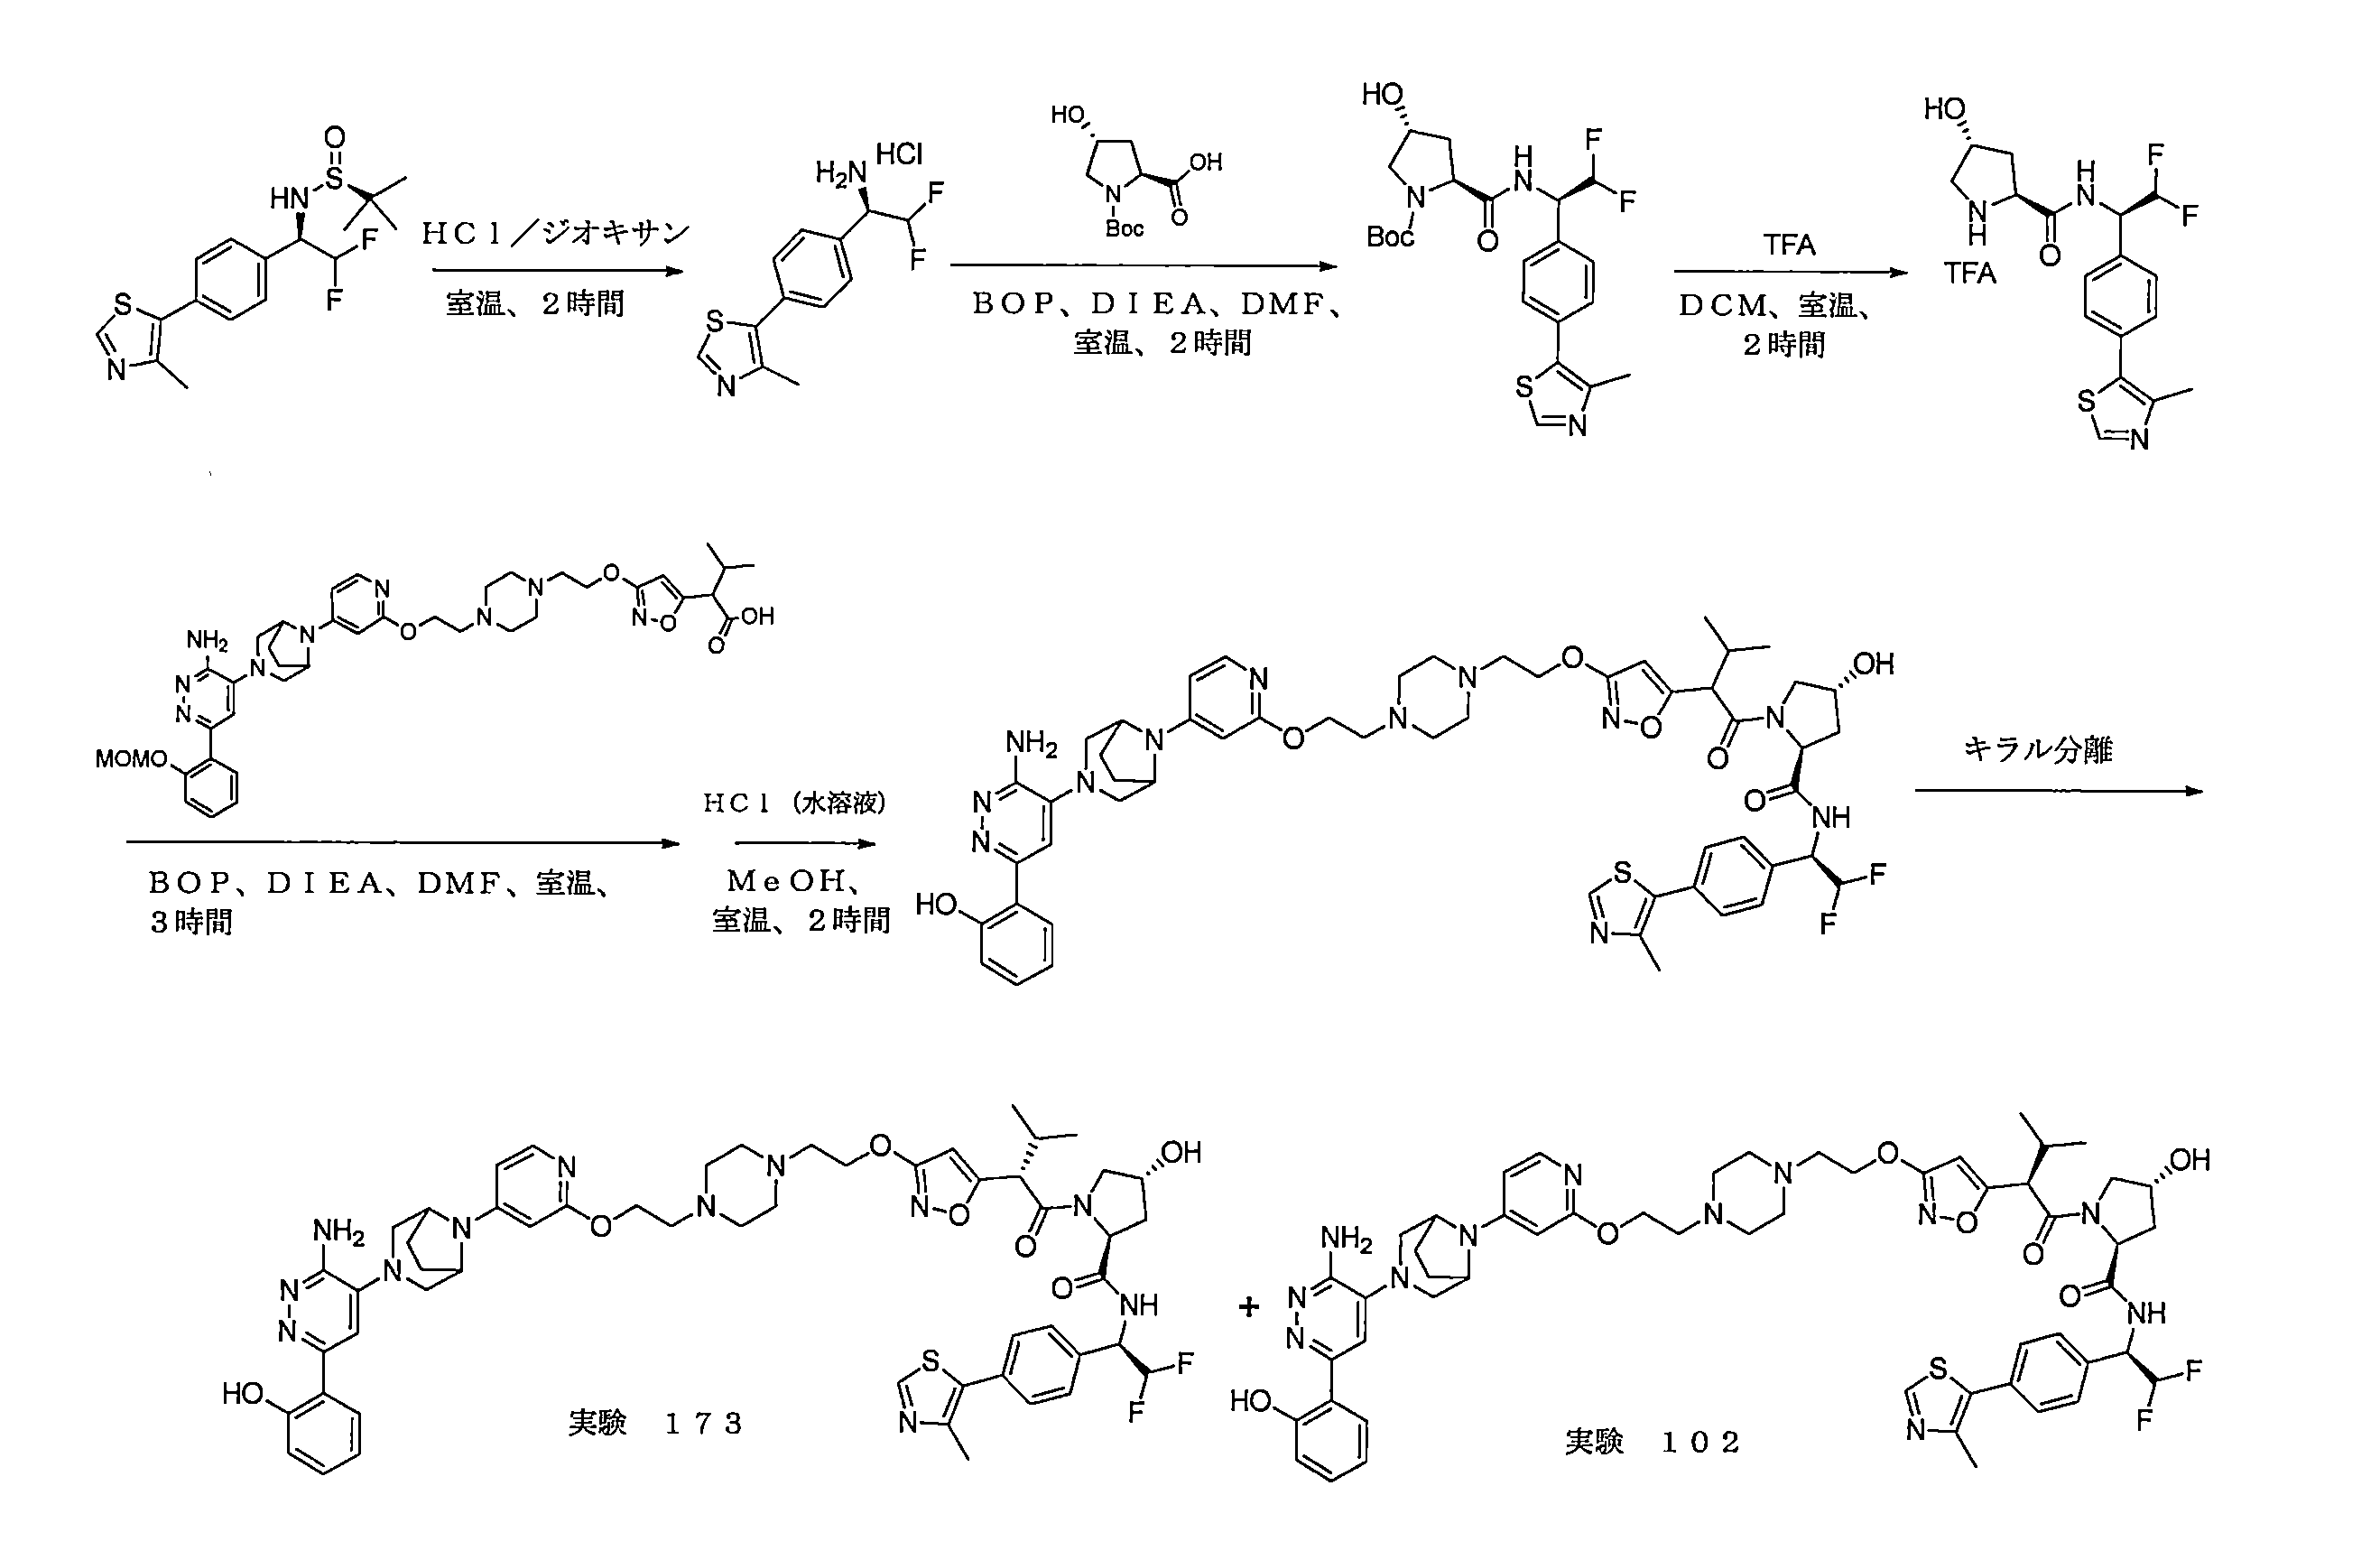
例示的な化合物267および268を、類似の手順を使用して調製した。 Exemplary compounds 267 and 268 were prepared using similar procedures.
例示的な化合物174の例示的な合成
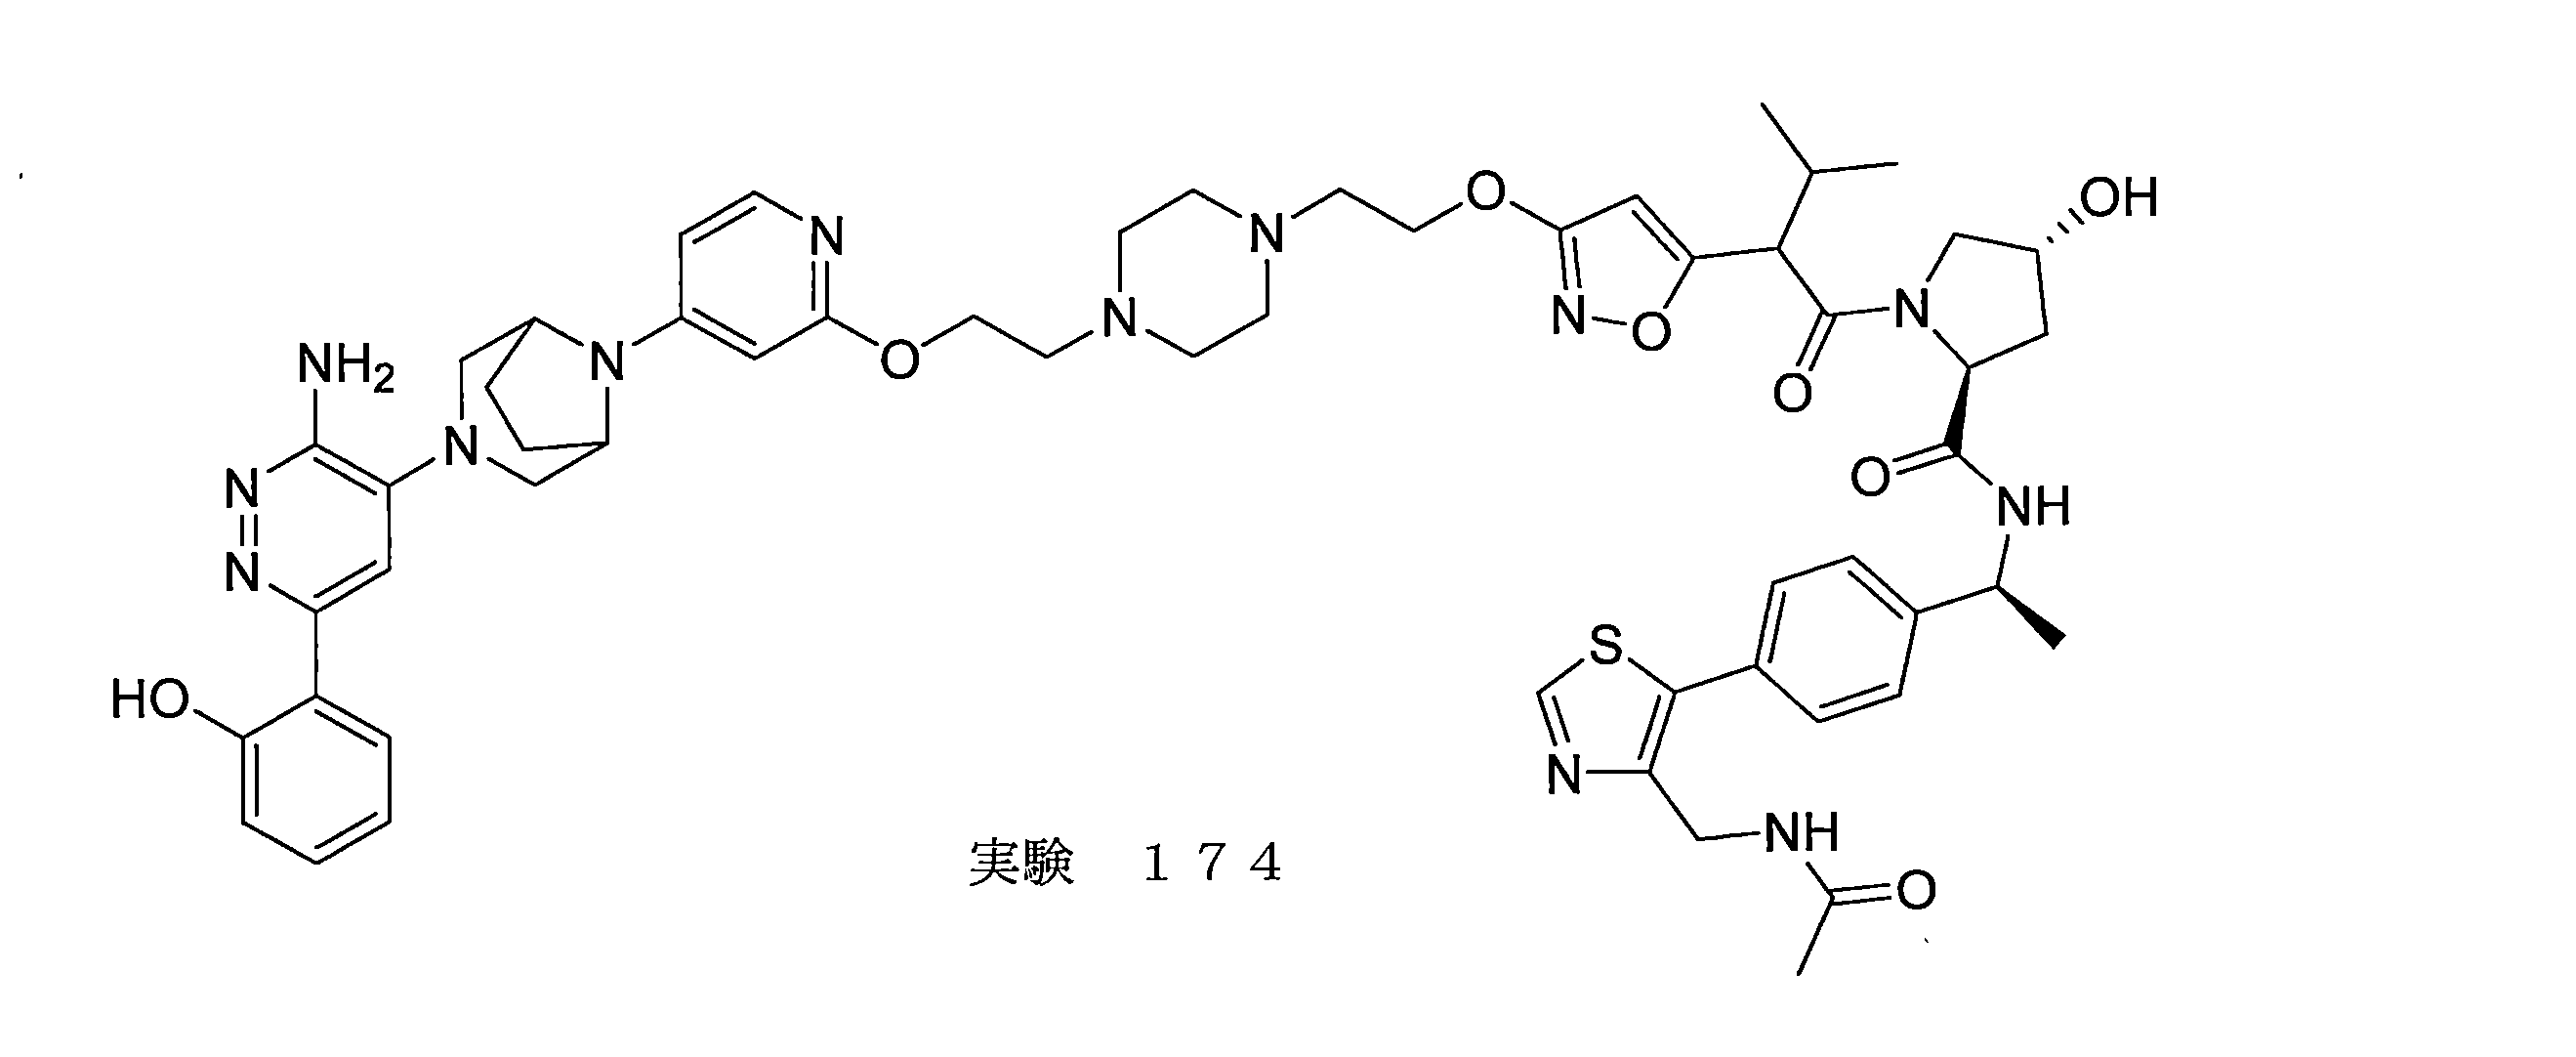
工程1

テトラヒドロフラン(8mL)中のtert-ブチルN-[(1S)-1-[4-[4-(ヒドロキシメチル)チアゾール-5-イル]フェニル]エチル]カルバメート(400mg、1.20mmol、1当量)、イソインドリン-1,3-ジオン(352mg、2.39mmol、2当量)、およびトリフェニルホスフィン(471mg、1.79mmol、1.5当量)の溶液に、N,N-ジイソプロピルアゾジカルボキシレート(290mg、1.44mmol、0.27mL、1.2当量)を0℃で添加した。混合物を15℃で16時間撹拌した。反応混合物を濃縮して、粗生成物を得、これを分取TLC(石油エーテル:酢酸エチル=1:1)によって精製して、tert-ブチルN-[(1S)-1-[4-[4-[(1,3-ジオキソイソインドリン-2-イル)メチル]チアゾール-5-イル]フェニル]エチル]カルバメート(560mg)を白色の固体として生成した。 To a solution of tert-butyl N-[(1S)-1-[4-[4-(hydroxymethyl)thiazol-5-yl]phenyl]ethyl]carbamate (400 mg, 1.20 mmol, 1 equiv.), isoindoline-1,3-dione (352 mg, 2.39 mmol, 2 equiv.), and triphenylphosphine (471 mg, 1.79 mmol, 1.5 equiv.) in tetrahydrofuran (8 mL) was added N,N-diisopropyl azodicarboxylate (290 mg, 1.44 mmol, 0.27 mL, 1.2 equiv.) at 0°C. The mixture was stirred at 15°C for 16 h. The reaction mixture was concentrated to give the crude product, which was purified by preparative TLC (petroleum ether: ethyl acetate = 1:1) to produce tert-butyl N-[(1S)-1-[4-[4-[(1,3-dioxoisoindolin-2-yl)methyl]thiazol-5-yl]phenyl]ethyl]carbamate (560 mg) as a white solid.
工程2
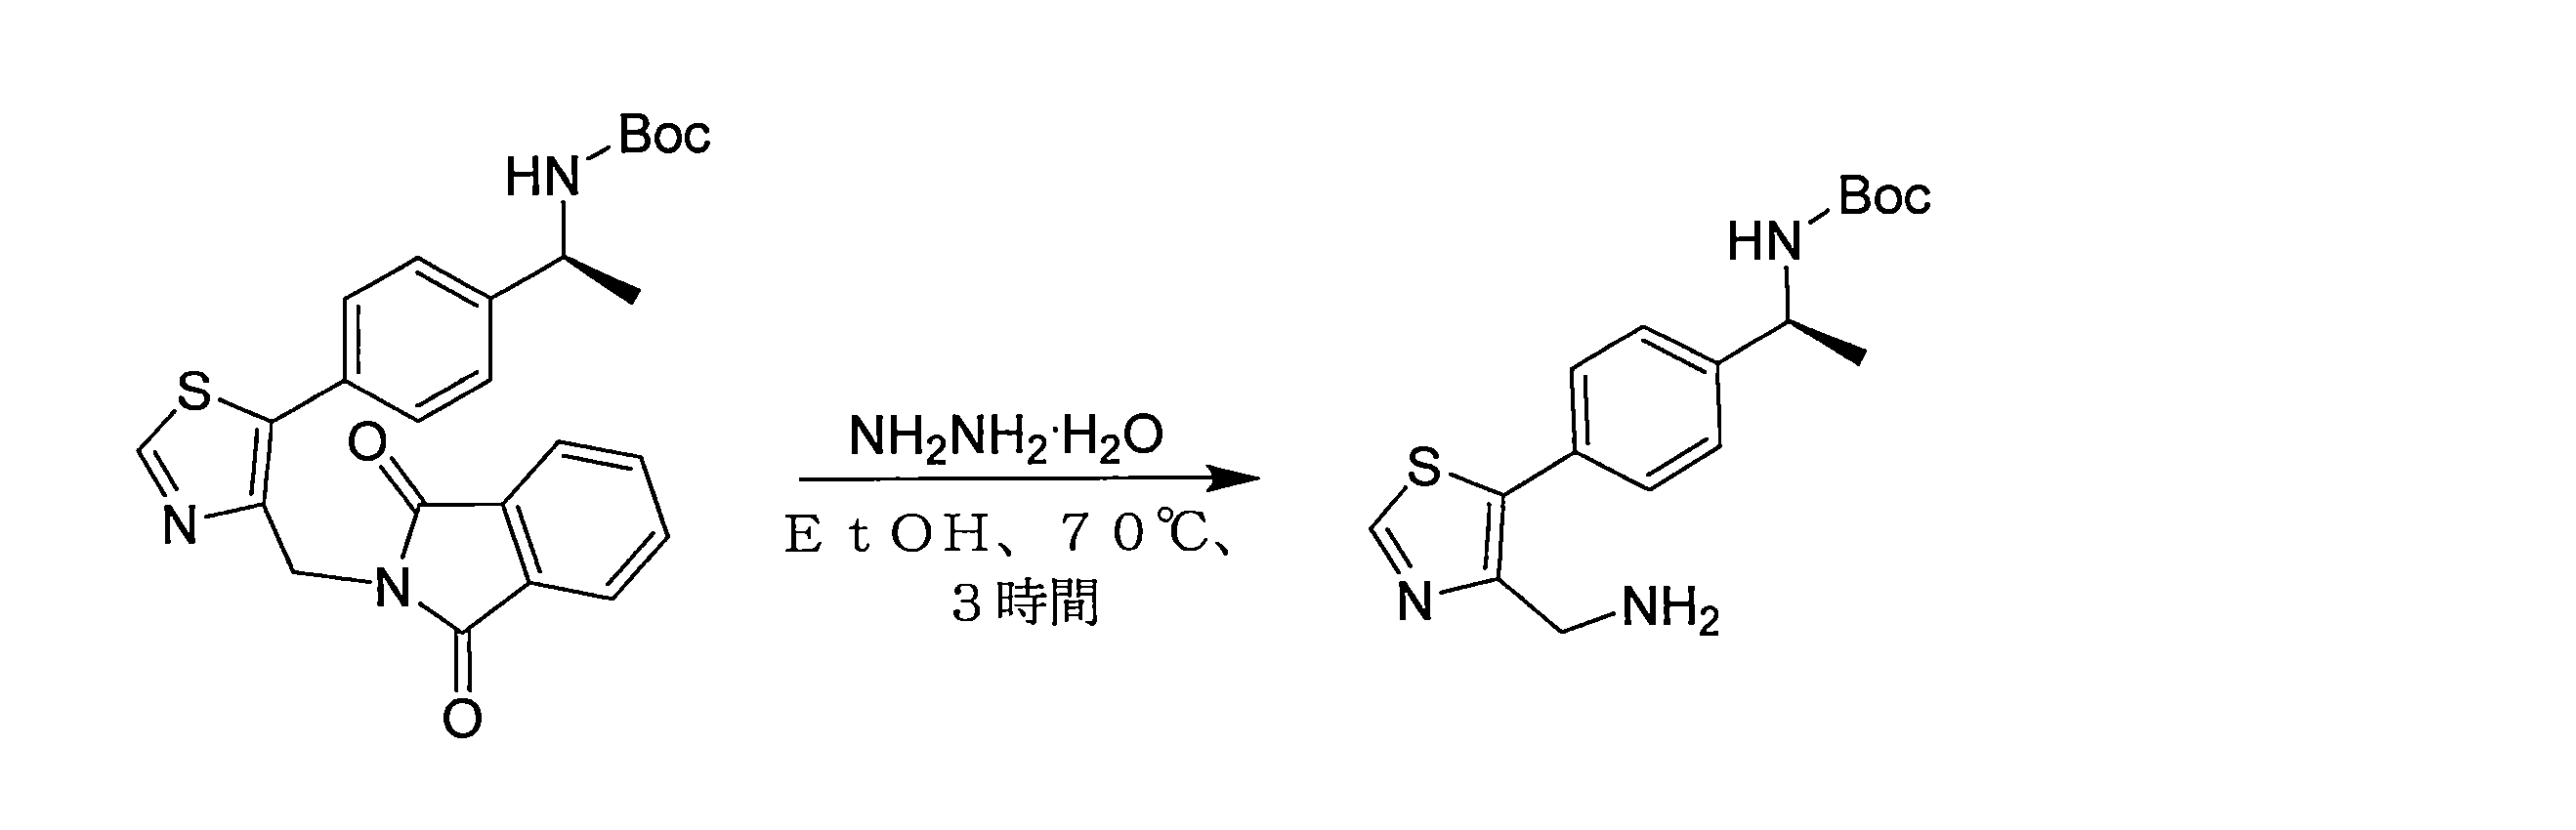
エタノール(24mL)中のtert-ブチルN-[(1S)-1-[4-[4-[(1,3-ジオキソイソインドリン-2-イル)メチル]チアゾール-5-イル]フェニル]エチル]カルバメート(600mg、1.29mmol、1当量)およびヒドラジン水和物(661mg、12.94mmol、0.64mL、純度98%、10当量)の混合物を、脱気し、窒素で3回パージし、次いで、混合物を窒素雰囲気下で、70℃で3時間撹拌した。反応混合物を濾過し、フィルターケーキをエタノール(30mL)で洗浄した。濾液を濃縮して粗生成物を得た。化合物tert-ブチルN-[(1S)-1-[4-[4-(アミノメチル)チアゾール-5-イル]フェニル]エチル]カルバメート(530mg、粗製)を白色の固体として得た。 A mixture of tert-butyl N-[(1S)-1-[4-[4-[(1,3-dioxoisoindolin-2-yl)methyl]thiazol-5-yl]phenyl]ethyl]carbamate (600 mg, 1.29 mmol, 1 eq.) and hydrazine hydrate (661 mg, 12.94 mmol, 0.64 mL, 98% purity, 10 eq.) in ethanol (24 mL) was degassed and purged with nitrogen three times, then the mixture was stirred under nitrogen atmosphere at 70°C for 3 h. The reaction mixture was filtered and the filter cake was washed with ethanol (30 mL). The filtrate was concentrated to give the crude product. Compound tert-butyl N-[(1S)-1-[4-[4-(aminomethyl)thiazol-5-yl]phenyl]ethyl]carbamate (530 mg, crude) was obtained as a white solid.
工程3
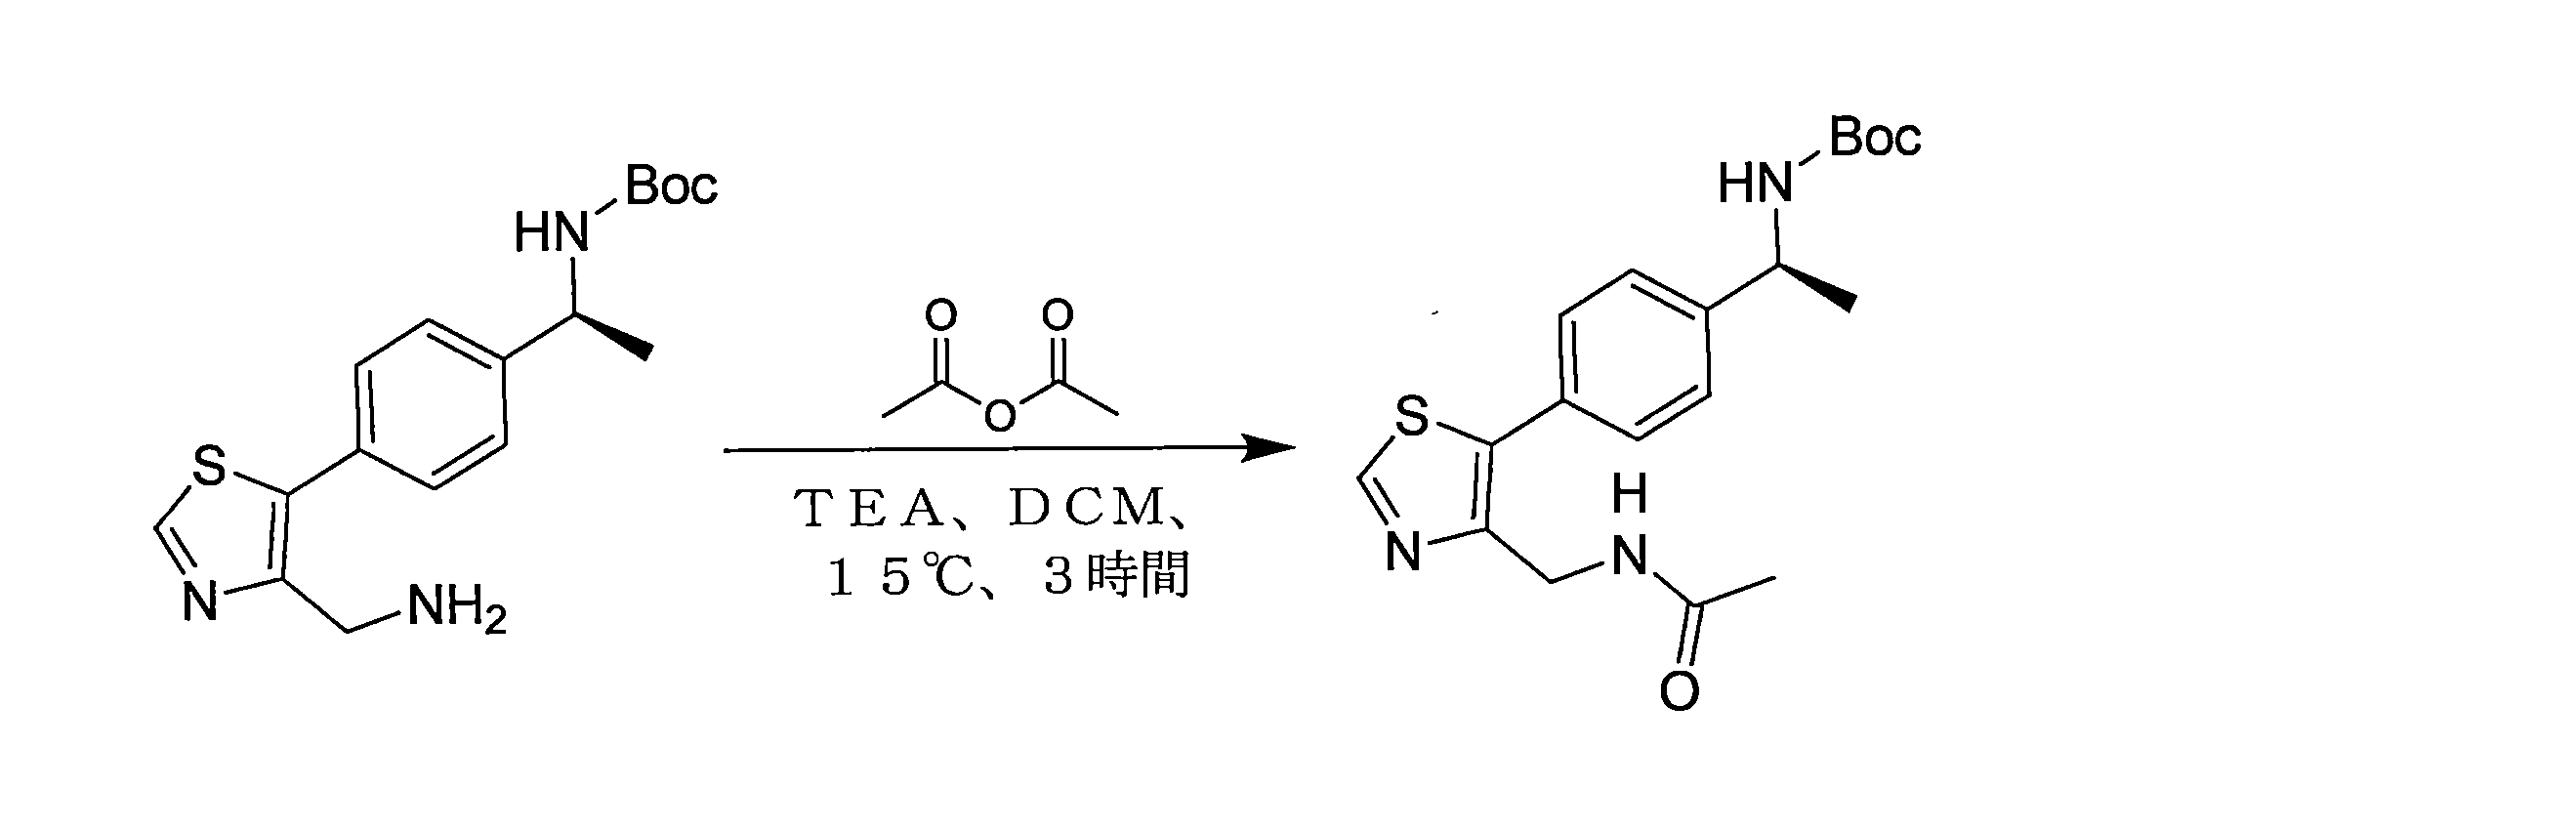
ジクロロメタン(30mL)中のtert-ブチルN-[(1S)-1-[4-[4-(アミノメチル)チアゾール-5-イル]フェニル]エチル]カルバメート(618mg、1.85mmol、1当量)およびトリエチルアミン(563mg、5.56mmol、0.77mL、3当量)の溶液に、無水酢酸(227mg、2.22mmol、0.21mL、1.2当量)を0℃で添加した。混合物を15℃で3時間撹拌した。反応混合物を水(30mL)で希釈し、次いで、有機層を濃縮して、粗生成物を得た。粗生成物を分取TLC(ジクロロメタン:メタノール=10:1)によって精製した。化合物tert-ブチルN-[(1S)-1-[4-[4-(アセトアミドメチル)チアゾール-5-イル]フェニル]エチル]カルバメート(400mg、1.07mmol、収率58%)を白色の固体として得た。 To a solution of tert-butyl N-[(1S)-1-[4-[4-(aminomethyl)thiazol-5-yl]phenyl]ethyl]carbamate (618 mg, 1.85 mmol, 1 eq.) and triethylamine (563 mg, 5.56 mmol, 0.77 mL, 3 eq.) in dichloromethane (30 mL), acetic anhydride (227 mg, 2.22 mmol, 0.21 mL, 1.2 eq.) was added at 0°C. The mixture was stirred at 15°C for 3 h. The reaction mixture was diluted with water (30 mL) and then the organic layer was concentrated to give the crude product. The crude product was purified by preparative TLC (dichloromethane:methanol=10:1). The compound tert-butyl N-[(1S)-1-[4-[4-(acetamidomethyl)thiazol-5-yl]phenyl]ethyl]carbamate (400 mg, 1.07 mmol, yield 58%) was obtained as a white solid.
例示的な化合物76について記載される手順を使用して、tert-ブチルN-[(1S)-1-[4-[4-(アセトアミドメチル)チアゾール-5-イル]フェニル]エチル]カルバメートを最終化合物に変換した。 Using the procedure described for exemplary compound 76, tert-butyl N-[(1S)-1-[4-[4-(acetamidomethyl)thiazol-5-yl]phenyl]ethyl]carbamate was converted to the final compound.
例示的な化合物175の例示的な合成
工程1

テトラヒドロフラン(50mL)中のtert-ブチルN-[(1S)-1-[4-(4-エチニルチアゾール-5-イル)フェニル]エチル]カルバメート(1.73g、5.27mmol、1当量)の溶液に、n-ブチルリチウム(2.5M、4.42mL、2.1当量)を窒素下で、-78℃で添加した。反応混合物を-50℃で2時間撹拌した。次いで、トリホルモール(474mg、5.27mmol、1当量)を添加し、反応混合物を22時間10℃に温めた。反応混合物を10℃でさらに20時間撹拌した。水(40mL)を添加し、混合物を酢酸エチル(40mL)で抽出した。有機層を硫酸ナトリウムで乾燥させ、次いで、真空下で濃縮した。残渣を、シリカゲルカラムクロマトグラフィー(石油エーテル中10~80%酢酸エチル)によって精製して、tert-ブチルN-[(1S)-1-[4-[4-(3-ヒドロキシプロパ-1-イニル)チアゾール-5-イル]フェニル]エチル]カルバメート(480mg、1.34mmol、収率25%)を白色の固体として得た。950mgの出発材料を回収した。 To a solution of tert-butyl N-[(1S)-1-[4-(4-ethynylthiazol-5-yl)phenyl]ethyl]carbamate (1.73 g, 5.27 mmol, 1 eq.) in tetrahydrofuran (50 mL) was added n-butyllithium (2.5 M, 4.42 mL, 2.1 eq.) under nitrogen at −78°C. The reaction mixture was stirred at −50°C for 2 h. Triformol (474 mg, 5.27 mmol, 1 eq.) was then added and the reaction mixture was warmed to 10°C for 22 h. The reaction mixture was stirred at 10°C for an additional 20 h. Water (40 mL) was added and the mixture was extracted with ethyl acetate (40 mL). The organic layer was dried over sodium sulfate and then concentrated under vacuum. The residue was purified by silica gel column chromatography (10-80% ethyl acetate in petroleum ether) to give tert-butyl N-[(1S)-1-[4-[4-(3-hydroxyprop-1-ynyl)thiazol-5-yl]phenyl]ethyl]carbamate (480 mg, 1.34 mmol, 25% yield) as a white solid. 950 mg of starting material was recovered.
例示的な化合物174について記載される手順を使用して、tert-ブチルN-[(1S)-1-[4-[4-(3-ヒドロキシプロパ-1-イニル)チアゾール-5-イル]フェニル]エチル]カルバメートを、tert-ブチル(S)-(1-(4-(4-(3-アセトアミドプロパ-1-イン-1-イル)チアゾール-5-イル)フェニル)エチル)カルバメートに変換した。

例示的な化合物76について記載されるものと類似の手順を使用して、tert-ブチル(S)-(1-(4-(4-(3-アセトアミドプロパ-1-イン-1-イル)チアゾール-5-イル)フェニル)エチル)カルバメートを最終化合物に変換した。 Using a procedure similar to that described for exemplary compound 76, tert-butyl (S)-(1-(4-(4-(3-acetamidoprop-1-yn-1-yl)thiazol-5-yl)phenyl)ethyl)carbamate was converted to the final compound.
例示的な化合物178の例示的な合成
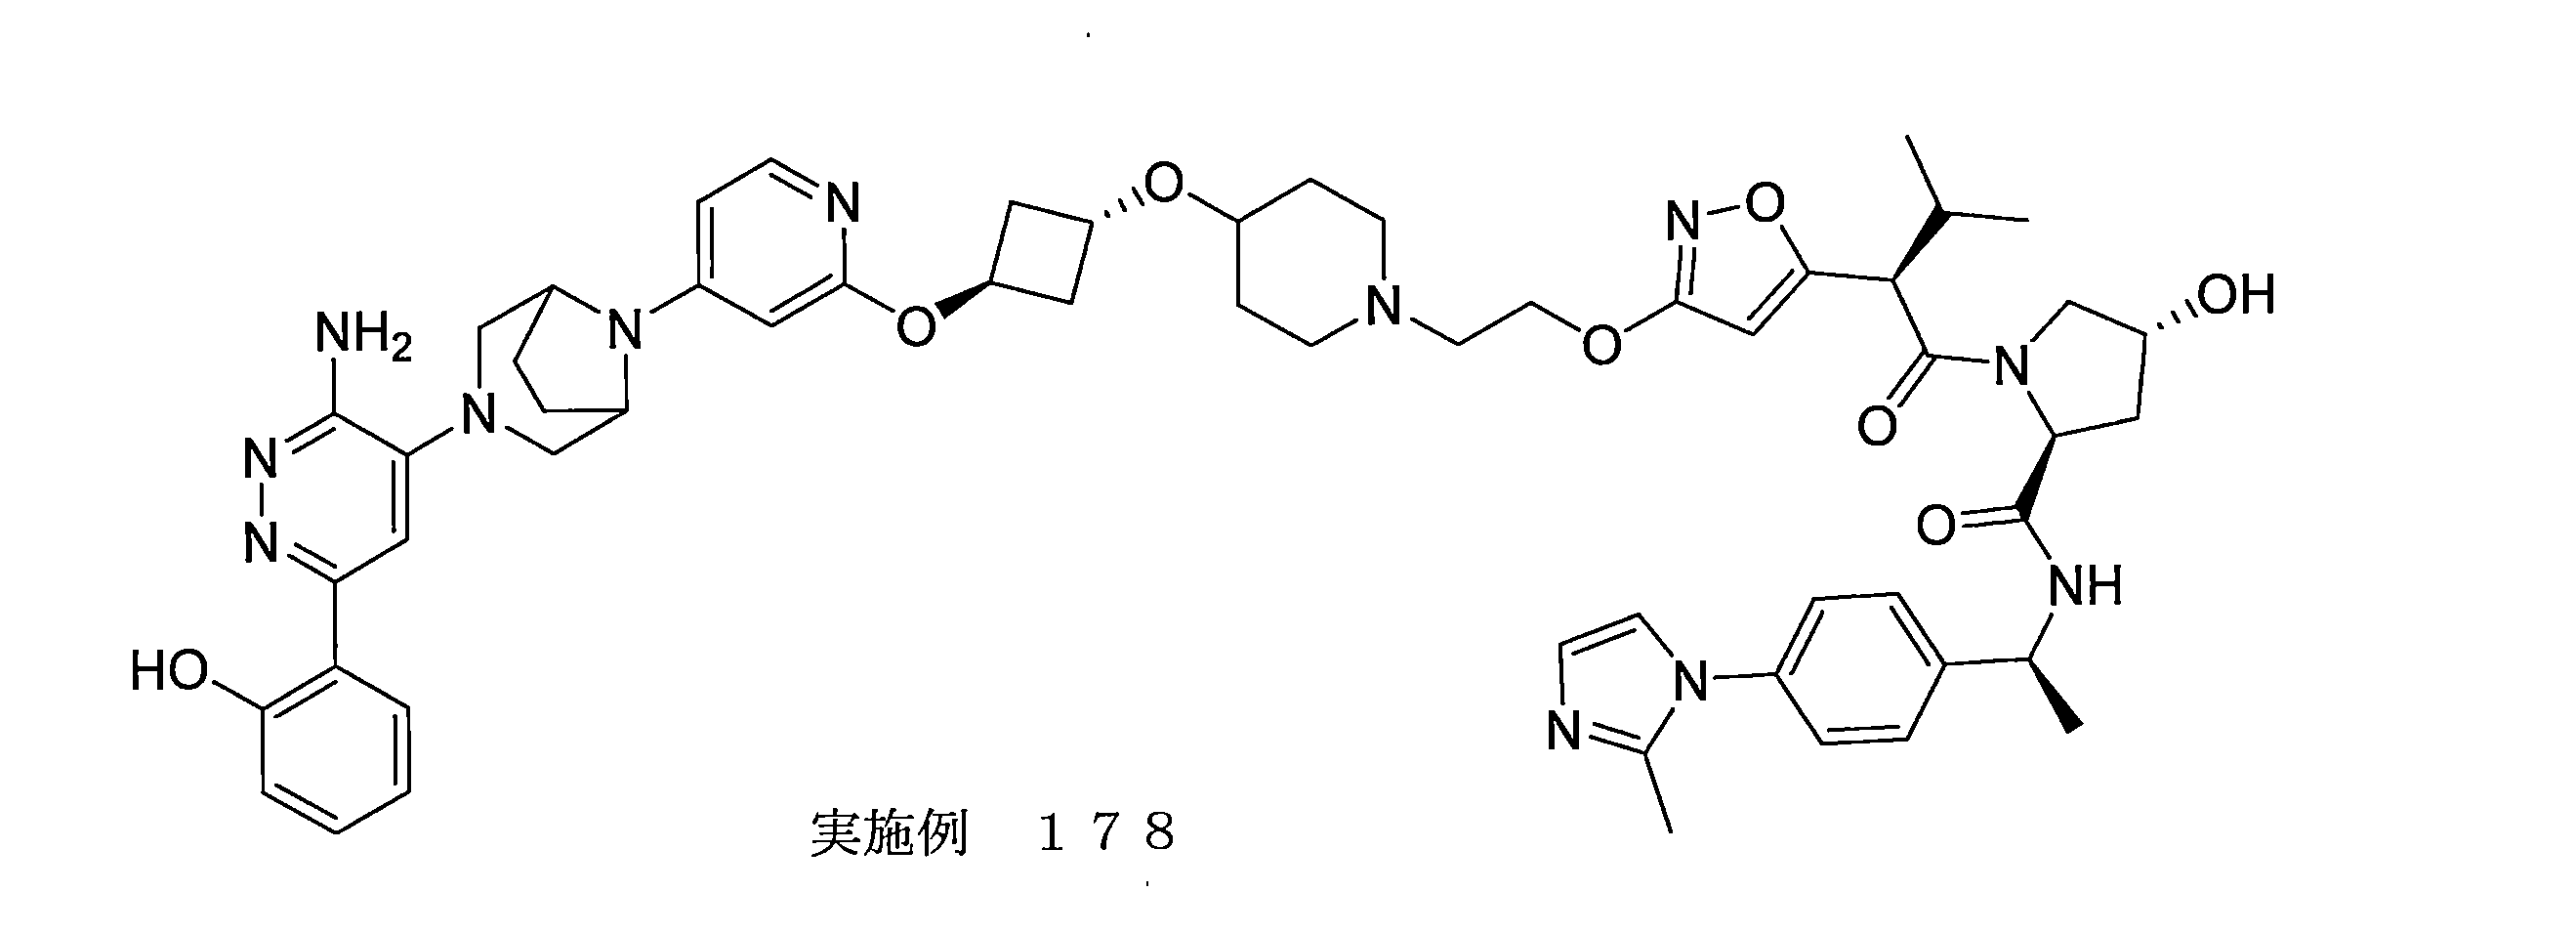
上に記載される手順を使用して、以下のスキームに従って調製した。
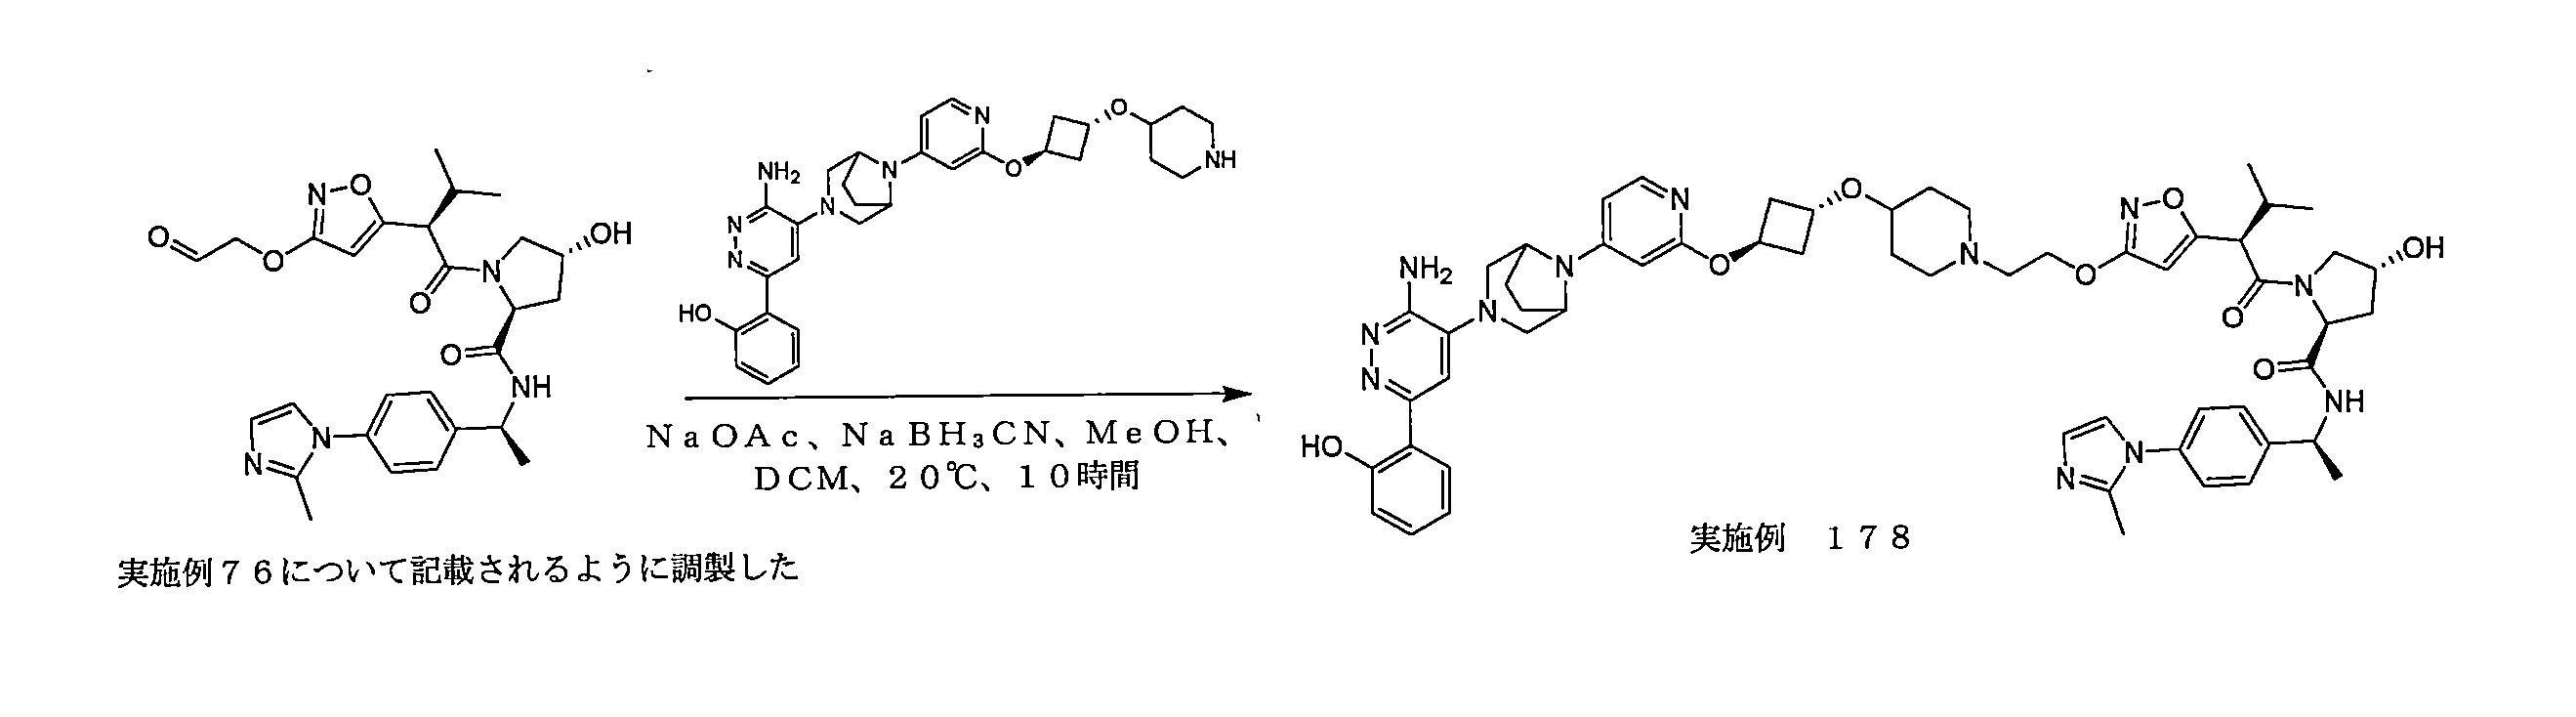
例示的な化合物176を、類似の手順を使用して調製した。 Exemplary compound 176 was prepared using a similar procedure.
例示的な化合物180の例示的な合成

上に記載される手順を使用して、以下のスキームに従って調製した。
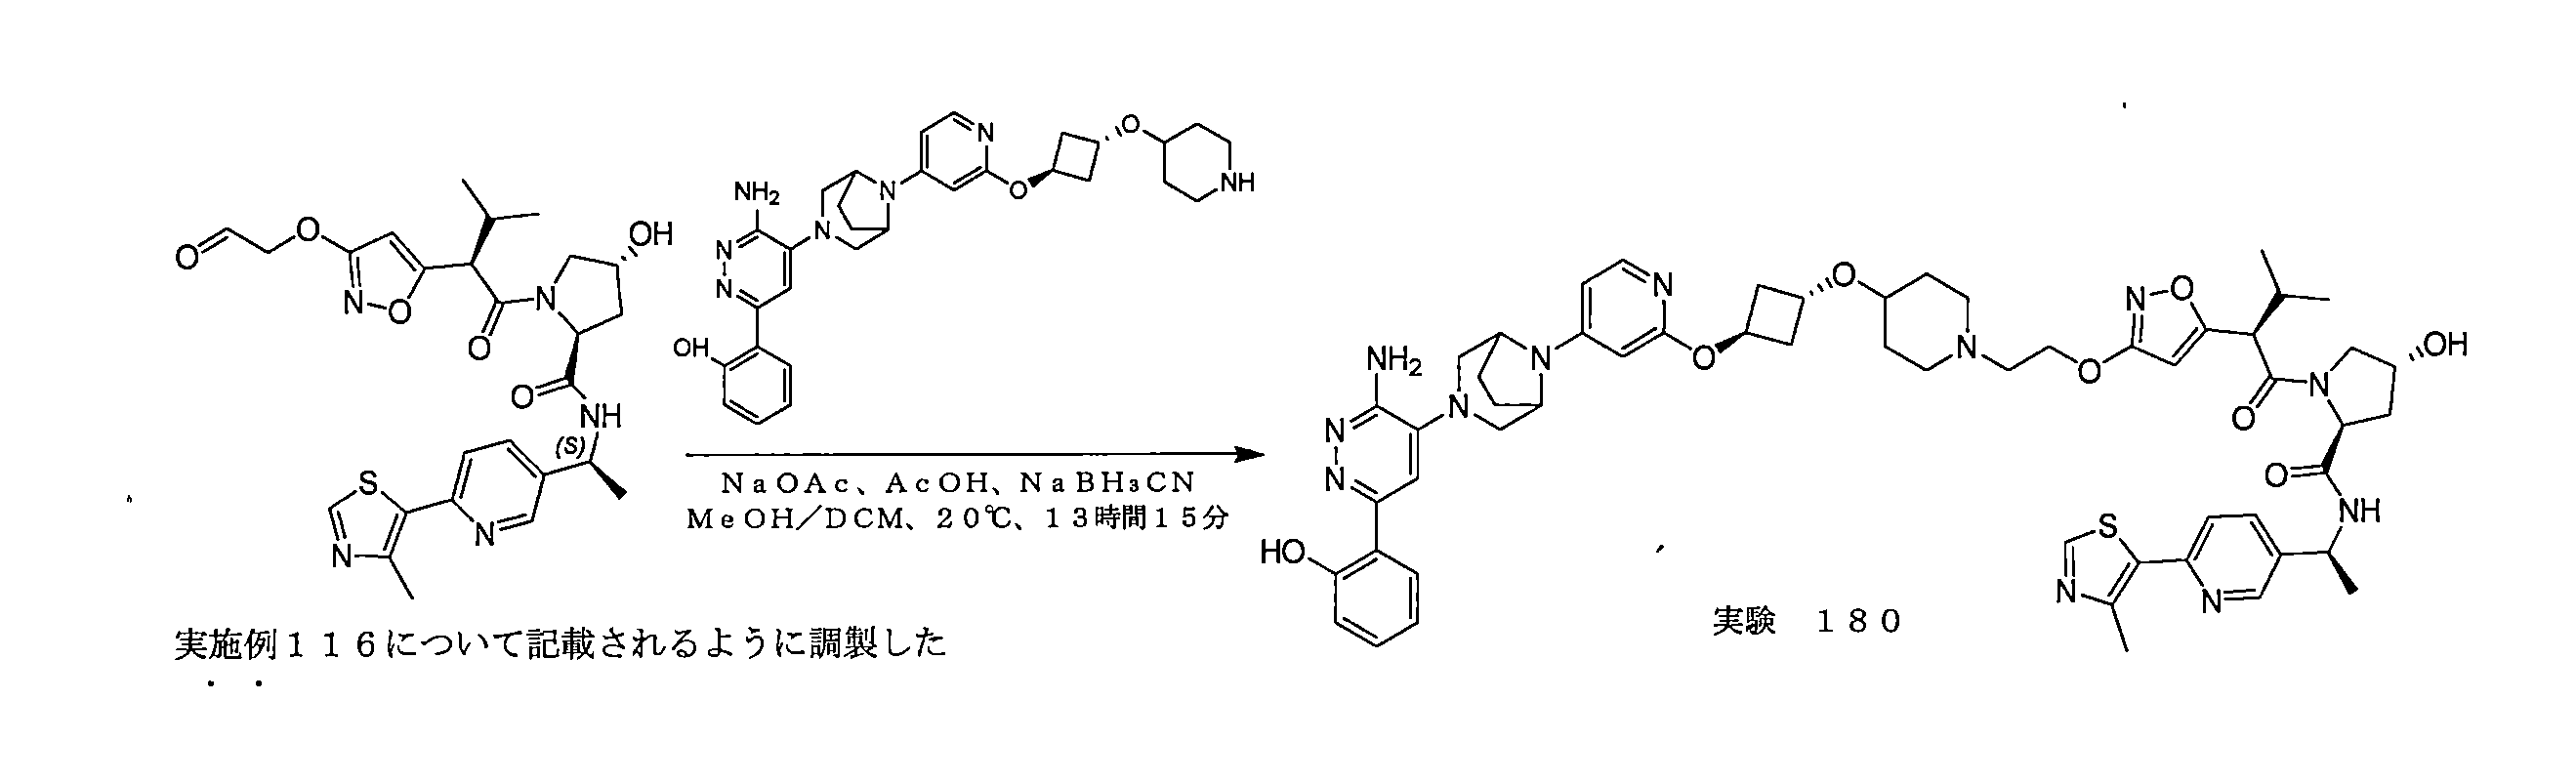
例示的な化合物179を、類似の手順を使用して調製した。 Exemplary compound 179 was prepared using a similar procedure.
例示的な化合物183の例示的な合成
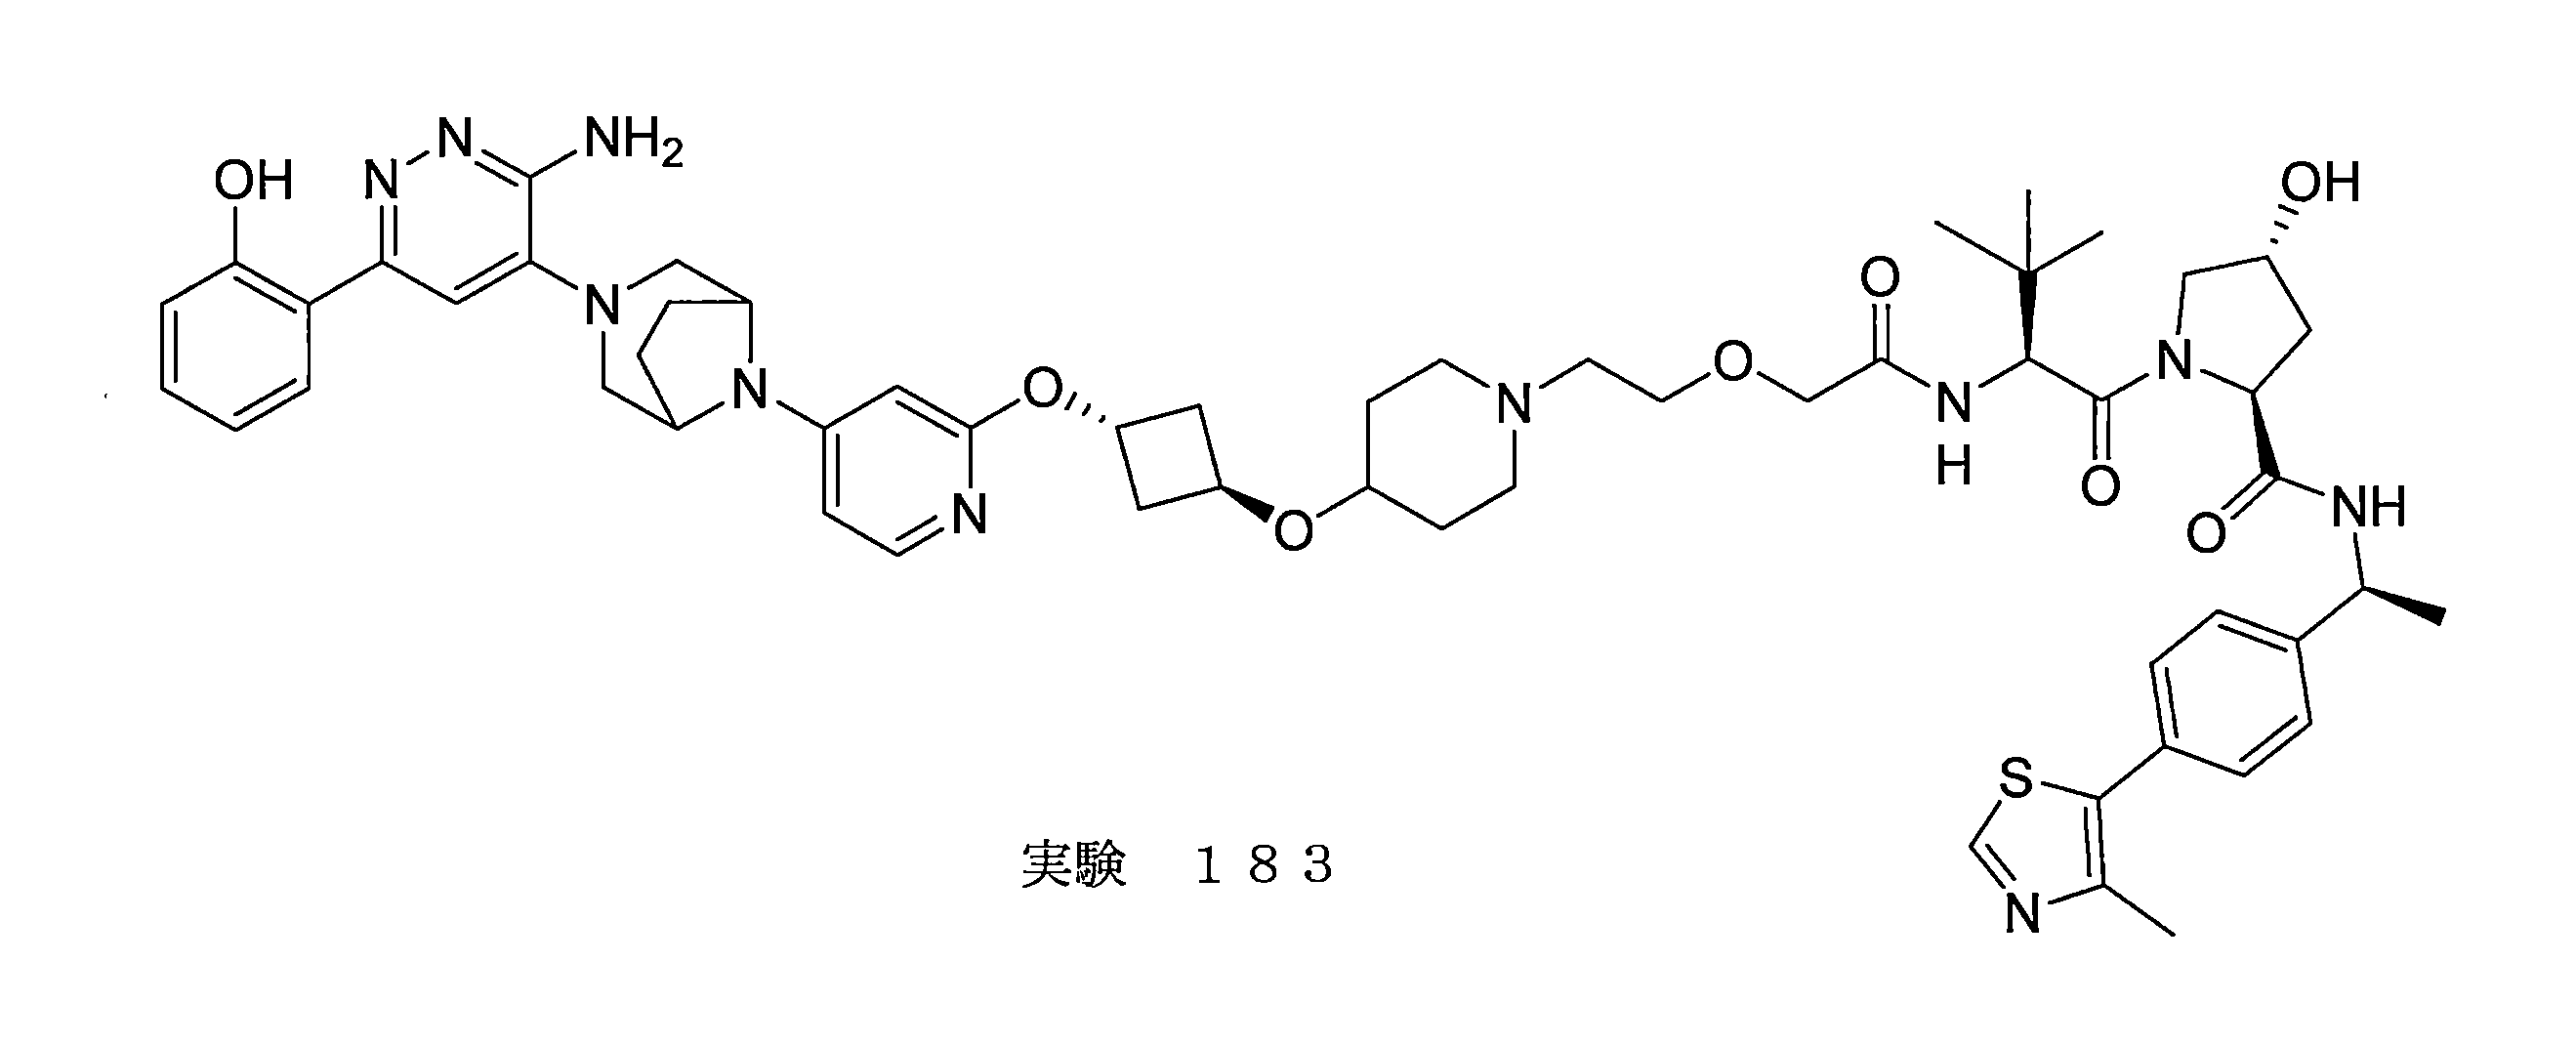
工程1

THF(50mL)中の2,2-ジエトキシエタノール(2.06g、15.38mmol、1.5当量)の溶液に、水素化ナトリウム(615mg、15.38mmol、純度60%、1.5当量)を0℃で添加し、混合物を0℃で30分間撹拌した。次いで、これに、THF(50mL)中のtert-ブチル2-ブロモ酢酸塩(2g、10.25mmol、1.52mL、1当量)の溶液を0℃で添加し、反応混合物を窒素下で、15℃で2時間撹拌した。混合物を0℃に冷却し、塩化アンモニウムの飽和水溶液によってクエンチした。水相を酢酸エチル(20mL×3)で抽出した。合わせた有機相をブライン(15mL×2)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、真空下で濃縮した。残渣を、シリカゲルクロマトグラフィー(石油エーテル/酢酸エチル=50/1~20/1)によって精製して、tert-ブチル2-(2,2-ジエトキシエトキシ)酢酸塩(883mg、3.56mmol、収率34%)を黄色の油として得た。 To a solution of 2,2-diethoxyethanol (2.06 g, 15.38 mmol, 1.5 eq.) in THF (50 mL) was added sodium hydride (615 mg, 15.38 mmol, 60% purity, 1.5 eq.) at 0°C and the mixture was stirred at 0°C for 30 minutes. To this was then added a solution of tert-butyl 2-bromoacetate (2 g, 10.25 mmol, 1.52 mL, 1 eq.) in THF (50 mL) at 0°C and the reaction mixture was stirred under nitrogen at 15°C for 2 hours. The mixture was cooled to 0°C and quenched by a saturated aqueous solution of ammonium chloride. The aqueous phase was extracted with ethyl acetate (20 mL x 3). The combined organic phase was washed with brine (15 mL x 2), dried over anhydrous sodium sulfate, filtered and concentrated under vacuum. The residue was purified by silica gel chromatography (petroleum ether/ethyl acetate = 50/1 to 20/1) to obtain tert-butyl 2-(2,2-diethoxyethoxy)acetate (883 mg, 3.56 mmol, 34% yield) as a yellow oil.
工程2

THF(3mL)、メタノール(3mL)、および水(6mL)の混合物中のtert-ブチル2-(2,2-ジエトキシエトキシ)酢酸塩(0.7g、2.82mmol、1当量)に、水酸化リチウム一水和物(1.18g、28.19mmol、10当量)を窒素下で、15℃で一度に添加した。混合物を2時間撹拌した。pHを1M塩酸で6に調整した。水相を酢酸エチル(35mL×4)で抽出した。合わせた有機層をブライン(20mL×2)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、真空下で濃縮させた。粗生成物2-(2,2-ジエトキシエトキシ)酢酸(460mg、粗製)を黄色の油として得た。 To tert-butyl 2-(2,2-diethoxyethoxy)acetate (0.7 g, 2.82 mmol, 1 equiv.) in a mixture of THF (3 mL), methanol (3 mL), and water (6 mL) was added lithium hydroxide monohydrate (1.18 g, 28.19 mmol, 10 equiv.) in one portion at 15°C under nitrogen. The mixture was stirred for 2 h. The pH was adjusted to 6 with 1 M hydrochloric acid. The aqueous phase was extracted with ethyl acetate (35 mL x 4). The combined organic layers were washed with brine (20 mL x 2), dried over anhydrous sodium sulfate, filtered, and concentrated under vacuum. The crude product 2-(2,2-diethoxyethoxy)acetic acid (460 mg, crude) was obtained as a yellow oil.
工程3

N,N-ジメチルホルムアミド(10mL)中の(2S,4R)-1-[(2S)-2-アミノ-3,3-ジメチル-ブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(0.2g、0.45mmol、1当量)および2-(2,2-ジエトキシエトキシ)酢酸(87mg、0.45mmol、1当量)の混合物に、O-(7-アザベンゾトリアゾール-1-イル)-N,N,N’,N’-テトラメチルウロニウムヘキサフルオロホスフェート(222mg、0.59mmol、1.3当量)およびトリエチルアミン(137mg、1.35mmol、0.19mL、3当量)を窒素下で、15℃で一度に添加した。混合物を15℃で1時間撹拌した。混合物を氷水(w/w=1/1、20mL)に注ぎ入れ、5分間撹拌した。水層を、酢酸エチル(20mL×4)で抽出した。合わせた有機相をブライン(15mL×2)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、真空中で濃縮した。残渣を、半分取逆相HPLC(ギ酸条件;カラム:Phenomenex Synergi C18 150*25*10um移動相:[水(0.225%FA)-ACN];B%:30%~60%、9分)によって精製した。生成物(2S,4R)-1-[(2S)-2-[[2-(2,2-ジエトキシエトキシ)アセチル]アミノ]-3,3-ジメチル-ブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(230mg、0.37mmol、収率82%)を黄色の油として得た。 To a mixture of (2S,4R)-1-[(2S)-2-amino-3,3-dimethyl-butanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (0.2 g, 0.45 mmol, 1 eq.) and 2-(2,2-diethoxyethoxy)acetic acid (87 mg, 0.45 mmol, 1 eq.) in N,N-dimethylformamide (10 mL), O-(7-azabenzotriazol-1-yl)-N,N,N',N'-tetramethyluronium hexafluorophosphate (222 mg, 0.59 mmol, 1.3 eq.) and triethylamine (137 mg, 1.35 mmol, 0.19 mL, 3 eq.) were added in one portion under nitrogen at 15°C. The mixture was stirred at 15°C for 1 h. The mixture was poured into ice water (w/w=1/1, 20 mL) and stirred for 5 min. The aqueous layer was extracted with ethyl acetate (20 mL×4). The combined organic phase was washed with brine (15 mL×2), dried over anhydrous sodium sulfate, filtered and concentrated in vacuo. The residue was purified by semi-preparative reverse phase HPLC (formic acid condition; column: Phenomenex Synergi C18 150*25*10um mobile phase: [water (0.225% FA)-ACN]; B%: 30%-60%, 9 min). The product (2S,4R)-1-[(2S)-2-[[2-(2,2-diethoxyethoxy)acetyl]amino]-3,3-dimethyl-butanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (230 mg, 0.37 mmol, 82% yield) was obtained as a yellow oil.
以下のスキームに従って、かつ他の実施例について上に記載される手順を使用して、(2S,4R)-1-[(2S)-2-[[2-(2,2-ジエトキシエトキシ)アセチル]アミノ]-3,3-ジメチル-ブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミドを最終化合物に変換した。
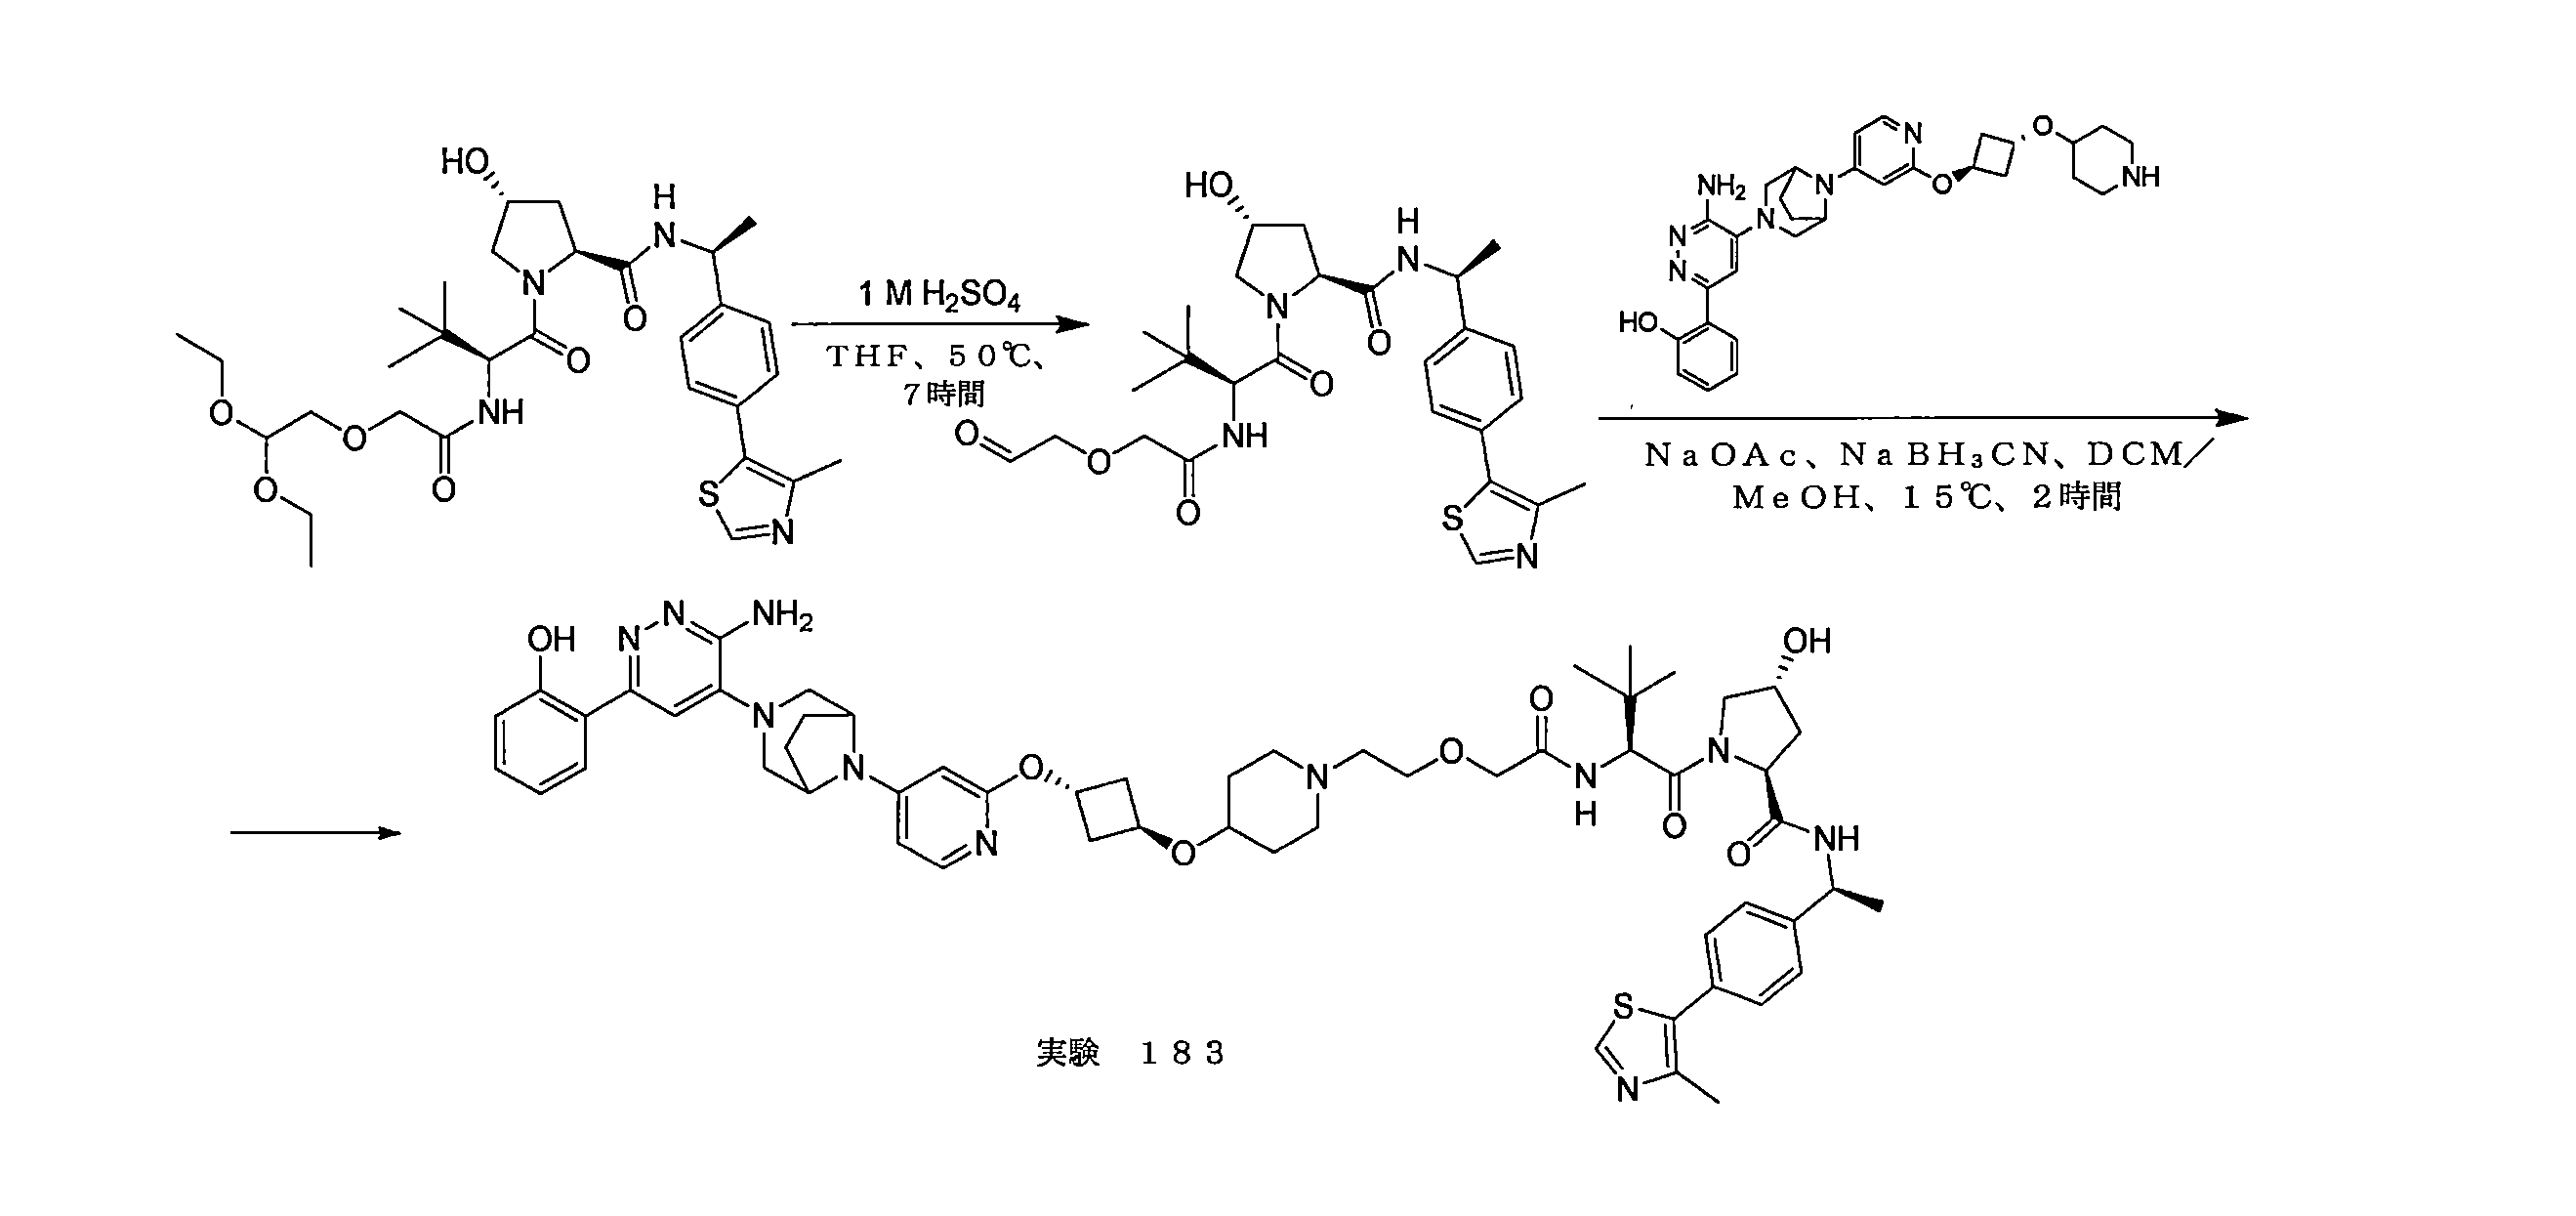
例示的な化合物86の例示的な合成

工程1

0℃で、THF(100mL)中のPPh3(4.68g、17.836mmol、3.0当量)の溶液に、DIAD(3.61g、17.836mmol、3.0当量)、および(1s,3s)-3-[[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]オキシ]シクロブタン-1-オール(3.0g、5.945mmol、1当量)を窒素雰囲気下で添加した。上記の混合物に、6-ブロモピリジン-3-オール(2.07g、11.891mmol、2.0当量)窒素雰囲気下で、0℃で5分間にわたって滴下で添加した。得られた混合物を、0℃で5分間撹拌し、次いで50℃に加熱し、50℃で2時間撹拌した。反応物を水(25mL)でクエンチし、得られた混合物をEtOAc(50mL×3)で抽出した。合わせた有機層をブライン(50mL)で洗浄し、無水硫酸ナトリウムで乾燥させ、減圧下で濃縮した。残渣を、CH2Cl2/MeOH(10:1)で溶出するシリカゲルカラムクロマトグラフィーによって精製して、6-[2-(メトキシメトキシ)フェニル]-4-(8-[2-[(1r,3r)-3-[(6-ブロモピリジン-3-イル)オキシ]シクロブトキシ]ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-イル)ピリダジン-3-アミン(2.3g、59%)を淡い黄色の固体として得た。 At 0 °C, to a solution of PPh3 (4.68 g, 17.836 mmol, 3.0 equiv.) in THF (100 mL) was added DIAD (3.61 g, 17.836 mmol, 3.0 equiv.), and (1s,3s)-3-[[4-(3-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl]oxy]cyclobutan-1-ol (3.0 g, 5.945 mmol, 1 equiv.) under nitrogen atmosphere. To the above mixture was added 6-bromopyridin-3-ol (2.07 g, 11.891 mmol, 2.0 equiv.) dropwise over 5 min at 0 °C under nitrogen atmosphere. The resulting mixture was stirred at 0°C for 5 minutes, then heated to 50°C and stirred at 50°C for 2 hours. The reaction was quenched with water (25 mL) and the resulting mixture was extracted with EtOAc (50 mL x 3). The combined organic layers were washed with brine (50 mL), dried over anhydrous sodium sulfate and concentrated under reduced pressure. The residue was purified by silica gel column chromatography eluting with CH2Cl2/MeOH (10:1) to give 6-[2-(methoxymethoxy)phenyl]-4-(8-[2-[(1r,3r)-3-[(6-bromopyridin-3-yl)oxy]cyclobutoxy]pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octan-3-yl)pyridazin-3-amine (2.3 g, 59%) as a pale yellow solid.
工程2

以下の反応を4つの並行の等しいバッチで行い、後処理および精製のために合わせた。各バッチにおいて、1,4-ジオキサン(15mL)および水(5mL)中の6-[2-(メトキシメトキシ)フェニル]-4-(8-[2-[(1r,3r)-3-[(6-ブロモピリジン-3-イル)オキシ]シクロブトキシ]ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-イル)ピリダジン-3-アミン(571mg、0.864mmol、1.00当量)およびtert-ブチルジメチル[[(2E)-3-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)プロパ-2-エン-1-イル]オキシ]シラン(387mg、1.297mmol、1.50当量)の溶液に、K2CO3(1.07g、0.008mmol、3.0当量)およびPd(dppf)Cl2-CH2Cl2(105mg、0.130mmol、0.15当量)を窒素雰囲気下で、室温で添加した。得られた混合物を窒素雰囲気下で、80℃で2時間撹拌した。各反応物を、室温で水(50mL)を添加することによってクエンチし、得られた混合物を合わせて、EtOAc(90mL×3)で抽出した。合わせた有機相をブラインで洗浄し、硫酸ナトリウムで乾燥させ、減圧下で濃縮した。残渣を、CH2Cl2/MeOH(10:1)で溶出するシリカゲルカラムクロマトグラフィーによって精製して、6-[2-(メトキシメトキシ)フェニル]-4-(8-[2-[(1r,3r)-3-([6-[(1E)-3-[(tert-ブチルジメチルシリル)オキシ]プロパ-1-エン-1-イル]ピリジン-3-イル]オキシ)シクロブトキシ]ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-イル)ピリダジン-3-アミン(合計2.1g、76%)を淡い黄色の固体として得た。 The following reaction was carried out in four parallel equal batches and combined for workup and purification. In each batch, 6-[2-(methoxymethoxy)phenyl]-4-(8-[2-[(1r,3r)-3-[(6-bromopyridin-3-yl)oxy]cyclobutoxy]pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octan-3-yl)pyridazin-3-amine (571 mg, 0.864 mmol, 1.00 equiv.) and tert-butyl ether in 1,4-dioxane (15 mL) and water (5 mL). To a solution of dimethyl[[(2E)-3-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)prop-2-en-1-yl]oxy]silane (387 mg, 1.297 mmol, 1.50 equiv.) was added K2CO3 (1.07 g, 0.008 mmol, 3.0 equiv.) and Pd(dppf)Cl2-CH2Cl2 (105 mg, 0.130 mmol, 0.15 equiv.) at room temperature under nitrogen atmosphere. The resulting mixture was stirred at 80 °C for 2 h under nitrogen atmosphere. Each reaction was quenched by adding water (50 mL) at room temperature and the resulting mixtures were combined and extracted with EtOAc (90 mL x 3). The combined organic phase was washed with brine, dried over sodium sulfate and concentrated under reduced pressure. The residue was purified by silica gel column chromatography eluting with CH2Cl2/MeOH (10:1) to give 6-[2-(methoxymethoxy)phenyl]-4-(8-[2-[(1r,3r)-3-([6-[(1E)-3-[(tert-butyldimethylsilyl)oxy]prop-1-en-1-yl]pyridin-3-yl]oxy)cyclobutoxy]pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octan-3-yl)pyridazin-3-amine (total 2.1 g, 76%) as a pale yellow solid.
工程3

THF(20mL)中の6-[2-(メトキシメトキシ)フェニル]-4-(8-[2-[(1r,3r)-3-([6-[(1E)-3-[(tert-ブチルジメチルシリル)オキシ]プロパ-1-エン-1-イル]ピリジン-3-イル]オキシ)シクロブトキシ]ピリジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-イル)ピリダジン-3-アミン(2.10g、2.79mmol、1.00当量)の撹拌溶液に、TBAF(1mol/L、8.3mL、8.3mmol、3.0当量)を窒素雰囲気下で、0℃で、滴下で添加した。得られた溶液を室温で2時間撹拌した。次いで、反応物を、水(25mL)を添加することによってクエンチした。得られた混合物を酢酸エチル(30mL×3)で抽出した。有機層を合わせ、無水硫酸ナトリウムで乾燥させ、濃縮した。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。これにより、3-[5-[(1r,3r)-3-[[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]オキシ]シクロブトキシ]ピリジン-2-イル]プロパ-2-エン-1-オール(1.1g、62%)を淡い黄色の固体として得た。 To a stirred solution of 6-[2-(methoxymethoxy)phenyl]-4-(8-[2-[(1r,3r)-3-([6-[(1E)-3-[(tert-butyldimethylsilyl)oxy]prop-1-en-1-yl]pyridin-3-yl]oxy)cyclobutoxy]pyridin-4-yl]-3,8-diazabicyclo[3.2.1]octan-3-yl)pyridazin-3-amine (2.10 g, 2.79 mmol, 1.00 equiv.) in THF (20 mL), TBAF (1 mol/L, 8.3 mL, 8.3 mmol, 3.0 equiv.) was added dropwise at 0° C. under a nitrogen atmosphere. The resulting solution was stirred at room temperature for 2 hours. The reaction was then quenched by adding water (25 mL). The resulting mixture was extracted with ethyl acetate (30 mL×3). The organic layers were combined, dried over anhydrous sodium sulfate and concentrated. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (10:1). This gave 3-[5-[(1r,3r)-3-[[4-(3-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl]oxy]cyclobutoxy]pyridin-2-yl]prop-2-en-1-ol (1.1 g, 62%) as a pale yellow solid.
工程4
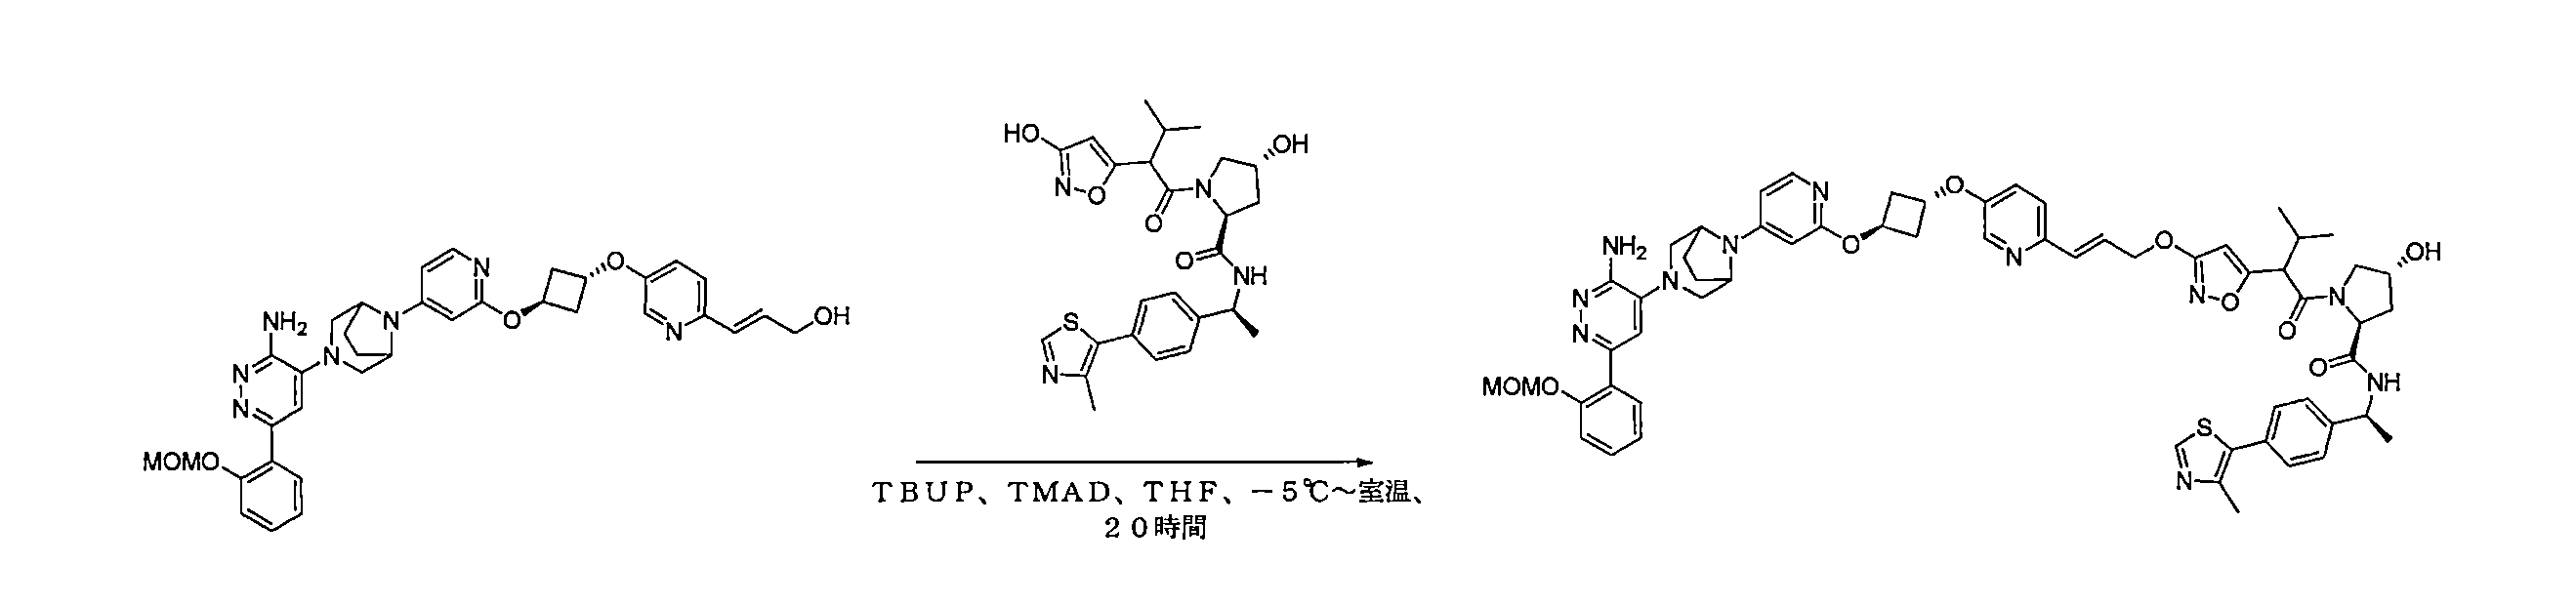
THF中のトリブチルホスフィン(634mg、3.14mmol、10.00当量)およびテトラメチルアゾジカルボキサミド(588mg、3.14mmol、10.00当量)の溶液に、(2S,4R)-4-ヒドロキシ-1-[2-(3-ヒドロキシ-1,2-オキサゾール-5-イル)-3-メチルブタノイル]-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(172mg、0.345mmol、1.10当量)および(2E)-3-[5-[(1r,3r)-3-[[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]オキシ]シクロブトキシ]ピリジン-2-イル]プロパ-2-エン-1-オール(70mg、0.11mmol、1当量)を窒素雰囲気下で、-10℃で添加した。得られた混合物を窒素雰囲気下で、室温で16時間撹拌した。反応物を、室温で水(50mL)を添加することによってクエンチし、得られた混合物をEtOAc(30mL×3)で抽出した。合わせた有機相をブライン(50mL)で洗浄し、硫酸ナトリウムで乾燥させ、減圧下で濃縮した。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラムクロマトグラフィーによって精製して、(2S,4R)-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]-1-[3-メチル-2-(3-[[(2E)-3-[5-[(1r,3r)-3-[[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]オキシ]シクロブトキシ]ピリジン-2-イル]プロパ-2-エン-1-イル]オキシ]-1,2-オキサゾール-5-イル)ブタノイル]ピロリジン-2-カルボキサミド(120mg、34%)を淡い黄色の固体として得た。 To a solution of tributylphosphine (634 mg, 3.14 mmol, 10.00 equiv.) and tetramethylazodicarboxamide (588 mg, 3.14 mmol, 10.00 equiv.) in THF, (2S,4R)-4-hydroxy-1-[2-(3-hydroxy-1,2-oxazol-5-yl)-3-methylbutanoyl]-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (172 mg, 0.345 mmol, 1.10 equiv.) and (2E)-3-[5-[(1r,3r)-3-[[4-(3-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl]oxy]cyclobutoxy]pyridin-2-yl]prop-2-en-1-ol (70 mg, 0.11 mmol, 1 equiv.) were added under nitrogen atmosphere at −10° C. The resulting mixture was stirred at room temperature under nitrogen atmosphere for 16 hours. The reaction was quenched by adding water (50 mL) at room temperature and the resulting mixture was extracted with EtOAc (30 mL×3). The combined organic phase was washed with brine (50 mL), dried over sodium sulfate and concentrated under reduced pressure. The residue was purified by silica gel column chromatography eluting with dichloromethane/methanol (10:1) to give (2S,4R)-4-hydroxy-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]-1-[3-methyl-2-(3-[[(2E)-3-[5-[(1r,3r)-3-[[4-(3-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl]oxy]cyclobutoxy]pyridin-2-yl]prop-2-en-1-yl]oxy]-1,2-oxazol-5-yl)butanoyl]pyrrolidine-2-carboxamide (120 mg, 34%) as a pale yellow solid.
他の実施例について上に記載される標準的な手順を使用して、(2S,4R)-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチル-1,3-チアゾール-5-イル)フェニル]エチル]-1-[3-メチル-2-(3-[[(2E)-3-[5-[(1r,3r)-3-[[4-(3-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル]オキシ]シクロブトキシ]ピリジン-2-イル]プロパ-2-エン-1-イル]オキシ]-1,2-オキサゾール-5-イル)ブタノイル]ピロリジン-2-カルボキサミドを最終化合物に変換した。 Using standard procedures described above for other examples, (2S,4R)-4-hydroxy-N-[(1S)-1-[4-(4-methyl-1,3-thiazol-5-yl)phenyl]ethyl]-1-[3-methyl-2-(3-[[(2E)-3-[5-[(1r,3r)-3-[[4-(3-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl]oxy]cyclobutoxy]pyridin-2-yl]prop-2-en-1-yl]oxy]-1,2-oxazol-5-yl)butanoyl]pyrrolidine-2-carboxamide was converted to the final compound.
例示的な化合物186、206、および207を、類似の手順を使用して調製した。 Exemplary compounds 186, 206, and 207 were prepared using similar procedures.
例示的な化合物189の例示的な合成
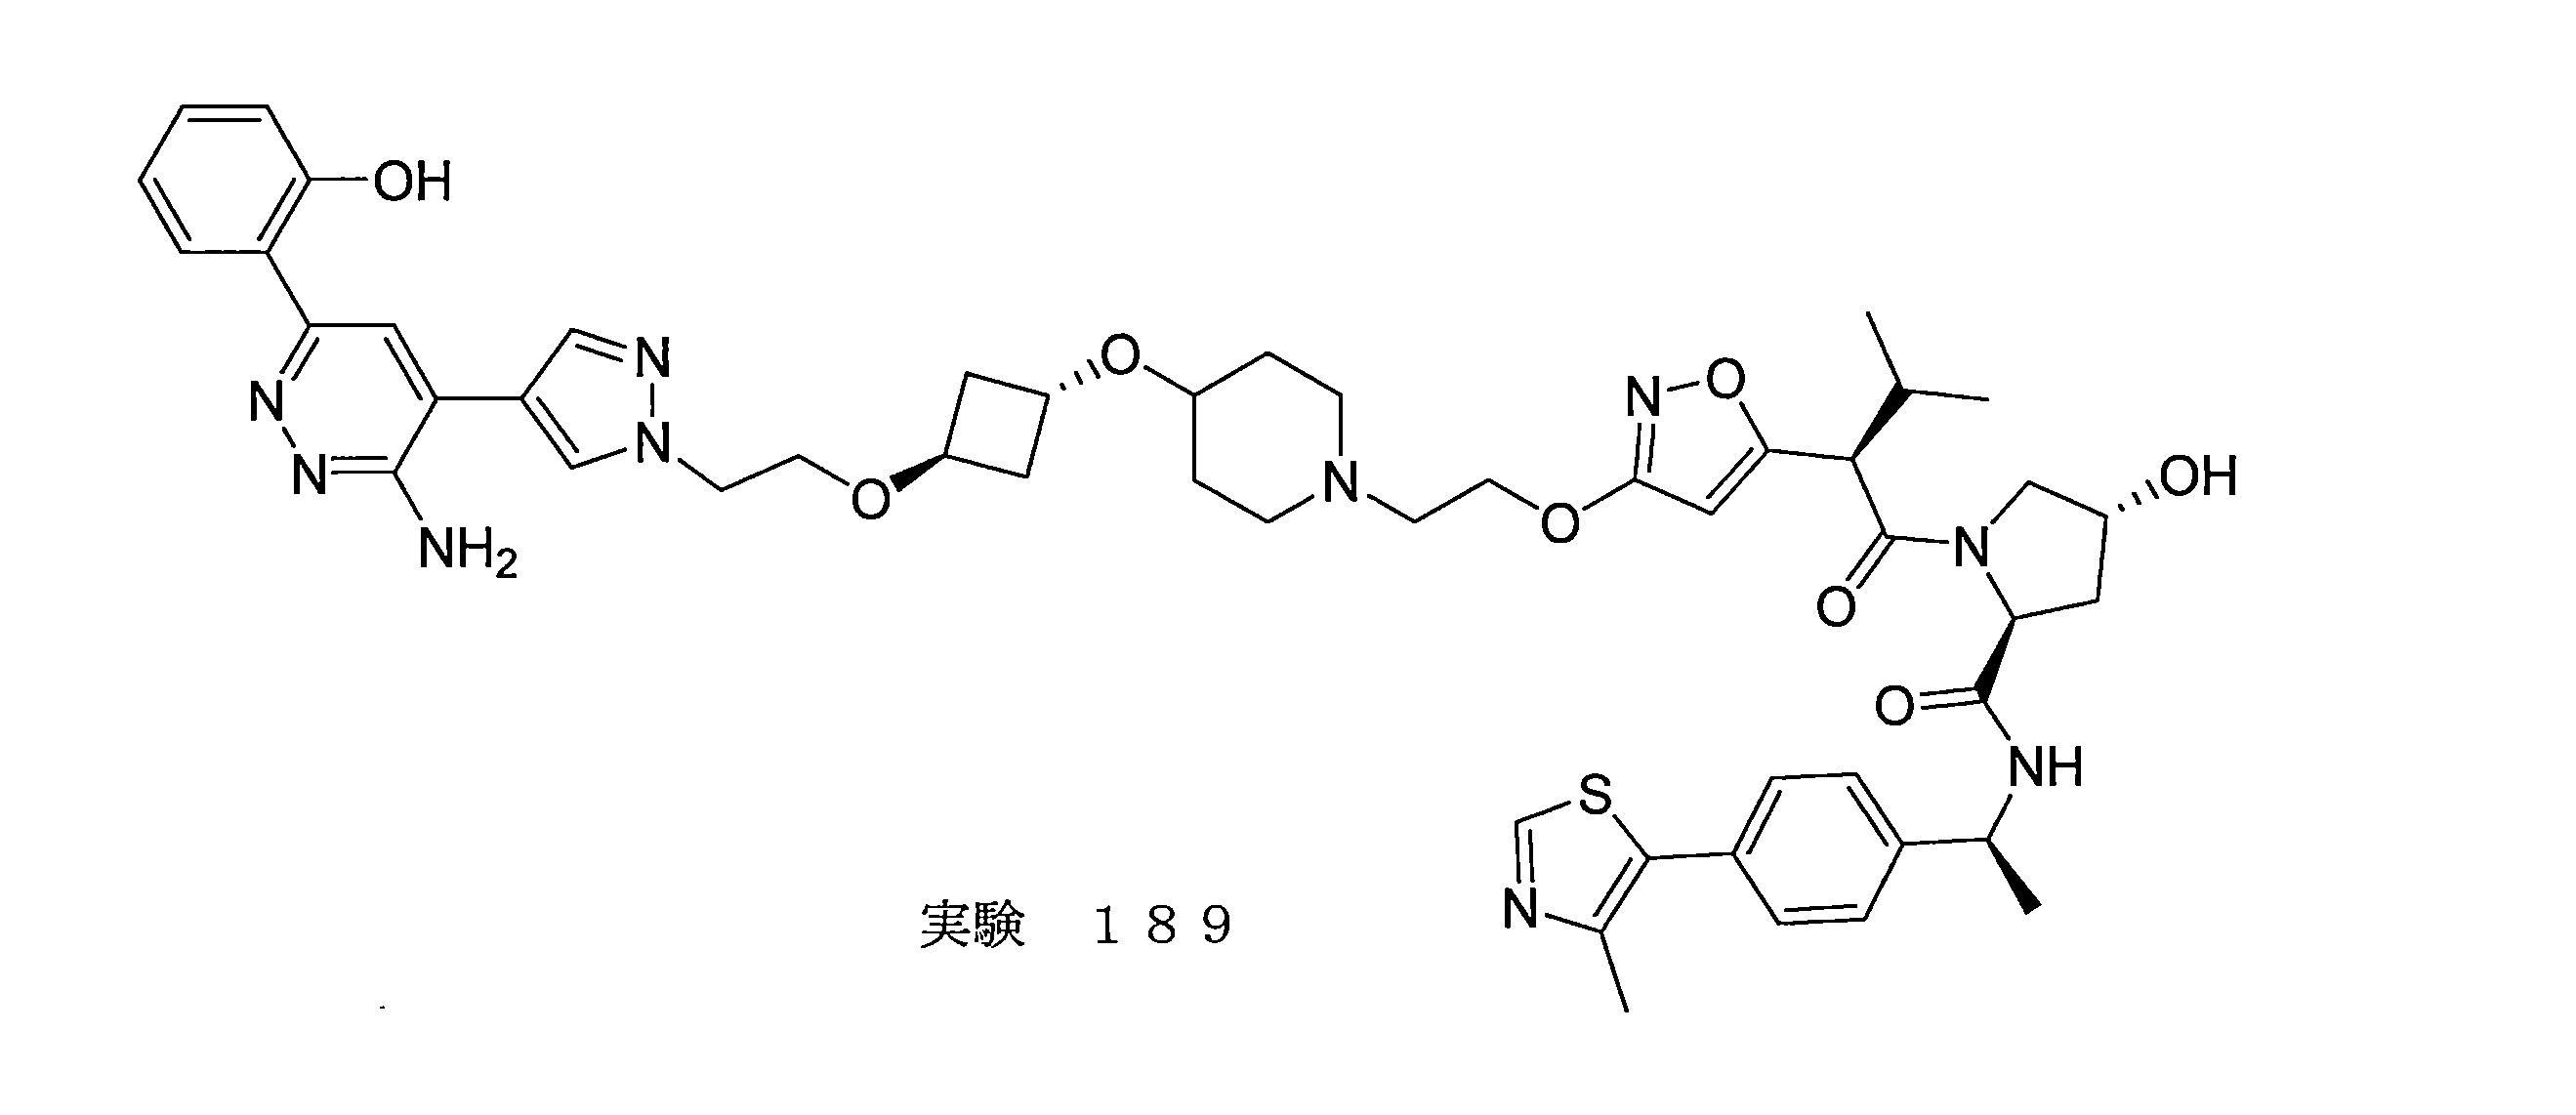
工程1
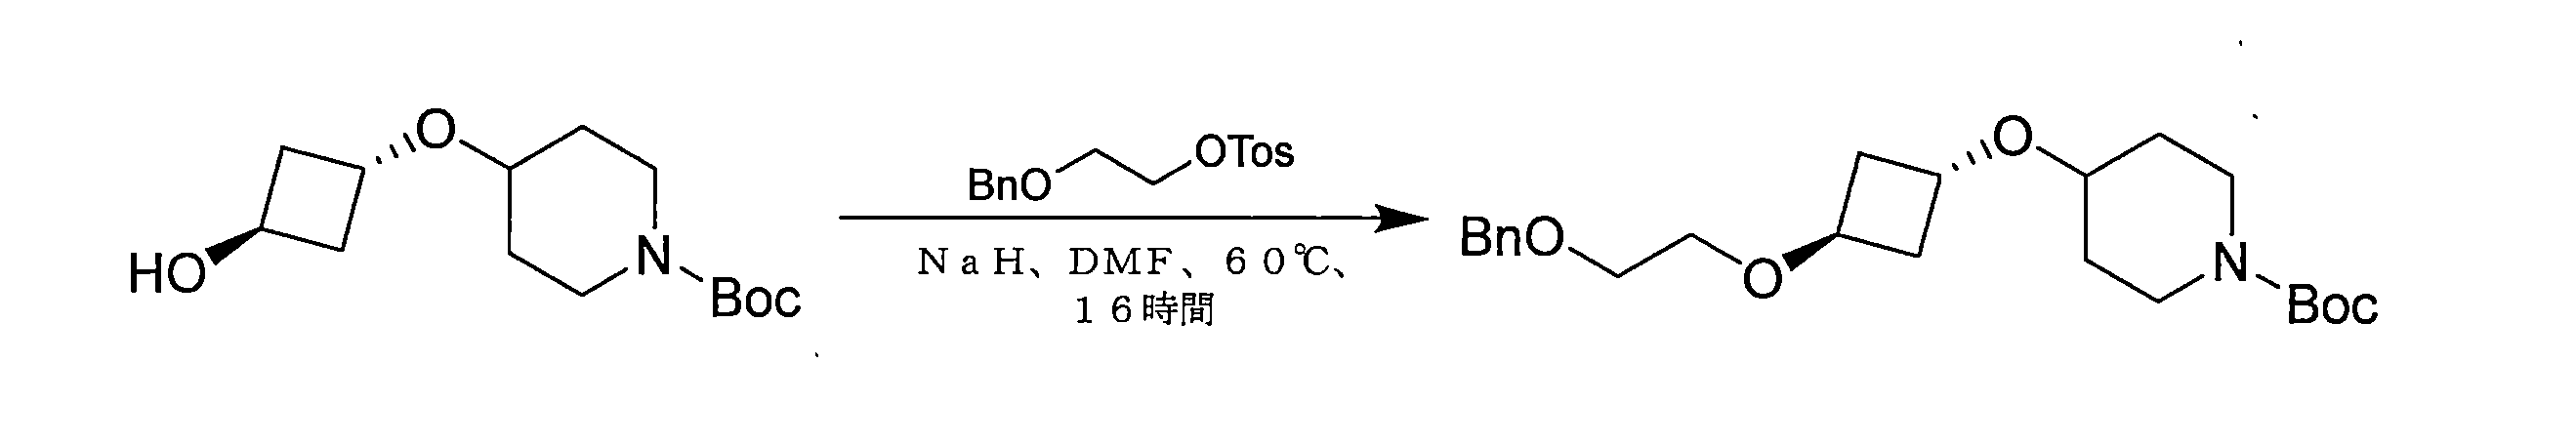
N,N-ジメチルホルムアミド(40mL)中のtert-ブチル4-(3-ヒドロキシシクロブトキシ)ピペリジン-1-カルボキシレート(2g、7.37mmol、1当量)の溶液に、水素化ナトリウム(353mg、8.84mmol、純度60%、1.2当量)を0℃で添加した。混合物を25℃で1時間撹拌した。次いで、N,N-ジメチルホルムアミド(2mL)中の2-ベンジルオキシエチル4-メチルベンゼンスルホネート(3.39g、11.06mmol、1.5当量)を添加した。混合物を60℃で15時間撹拌した。反応混合物を、水(200mL)を添加することによってクエンチし、酢酸エチル(200mL×3)で抽出した。合わせた有機層をブライン(200mL×3)で洗浄し、無水硫酸ナトリウム上で乾燥させ、濾過し、減圧下で濃縮して残渣を得た。残渣を、分取HPLC(カラム:Phenomenex Synergi Max-RP 250*50mm*10um;移動相:[水(0.225%FA)-ACN];B%:50ACN%~80ACN%、26分、20%分)によって精製した。化合物tert-ブチル4-[3-(2-ベンジルオキシエトキシ)シクロブトキシ]ピペリジン-1-カルボキシレート(2.1g、5.18mmol、収率70%)を無色の油として得た。 To a solution of tert-butyl 4-(3-hydroxycyclobutoxy)piperidine-1-carboxylate (2 g, 7.37 mmol, 1 eq.) in N,N-dimethylformamide (40 mL) was added sodium hydride (353 mg, 8.84 mmol, 60% purity, 1.2 eq.) at 0°C. The mixture was stirred at 25°C for 1 h. Then, 2-benzyloxyethyl 4-methylbenzenesulfonate (3.39 g, 11.06 mmol, 1.5 eq.) in N,N-dimethylformamide (2 mL) was added. The mixture was stirred at 60°C for 15 h. The reaction mixture was quenched by adding water (200 mL) and extracted with ethyl acetate (200 mL x 3). The combined organic layers were washed with brine (200 mL x 3), dried over anhydrous sodium sulfate, filtered, and concentrated under reduced pressure to give a residue. The residue was purified by preparative HPLC (column: Phenomenex Synergi Max-RP 250*50mm*10um; mobile phase: [water (0.225% FA)-ACN]; B%: 50ACN%-80ACN%, 26 min, 20% min). The compound tert-butyl 4-[3-(2-benzyloxyethoxy)cyclobutoxy]piperidine-1-carboxylate (2.1 g, 5.18 mmol, 70% yield) was obtained as a colorless oil.
工程2

エタノール(20mL)およびテトラヒドロフラン(20mL)中のtert-ブチル4-[3-(2-ベンイジルオキシエトキシ)シクロブトキシ]ピペリジン-1-カルボキシレート(2.1g、5.18mmol、1当量)の溶液に、5%パラジウム活性炭素触媒(500mg)を窒素下で添加した。懸濁液を真空下で脱気し、水素で3回パージした。混合物を、水素(15Psi)下で、25℃で16時間撹拌した。反応混合物を減圧下で濾過および濃縮し、残渣を得た。化合物tert-ブチル4-[3-(2-ヒドロキシエトキシ)シクロブトキシ]ピペリジン-1-カルボキシレート(1.6g、5.07mmol、収率97%)を無色の油として得た。 To a solution of tert-butyl 4-[3-(2-benzyloxyethoxy)cyclobutoxy]piperidine-1-carboxylate (2.1 g, 5.18 mmol, 1 equiv.) in ethanol (20 mL) and tetrahydrofuran (20 mL) was added 5% palladium on activated carbon catalyst (500 mg) under nitrogen. The suspension was degassed under vacuum and purged with hydrogen three times. The mixture was stirred under hydrogen (15 Psi) at 25°C for 16 h. The reaction mixture was filtered and concentrated under reduced pressure to give a residue. The compound tert-butyl 4-[3-(2-hydroxyethoxy)cyclobutoxy]piperidine-1-carboxylate (1.6 g, 5.07 mmol, 97% yield) was obtained as a colorless oil.
他の実施例について上に記載される手順を使用して、以下のスキームに従って、tert-ブチル4-[3-(2-ヒドロキシエトキシ)シクロブトキシ]ピペリジン-1-カルボキシレートを最終化合物に変換した。
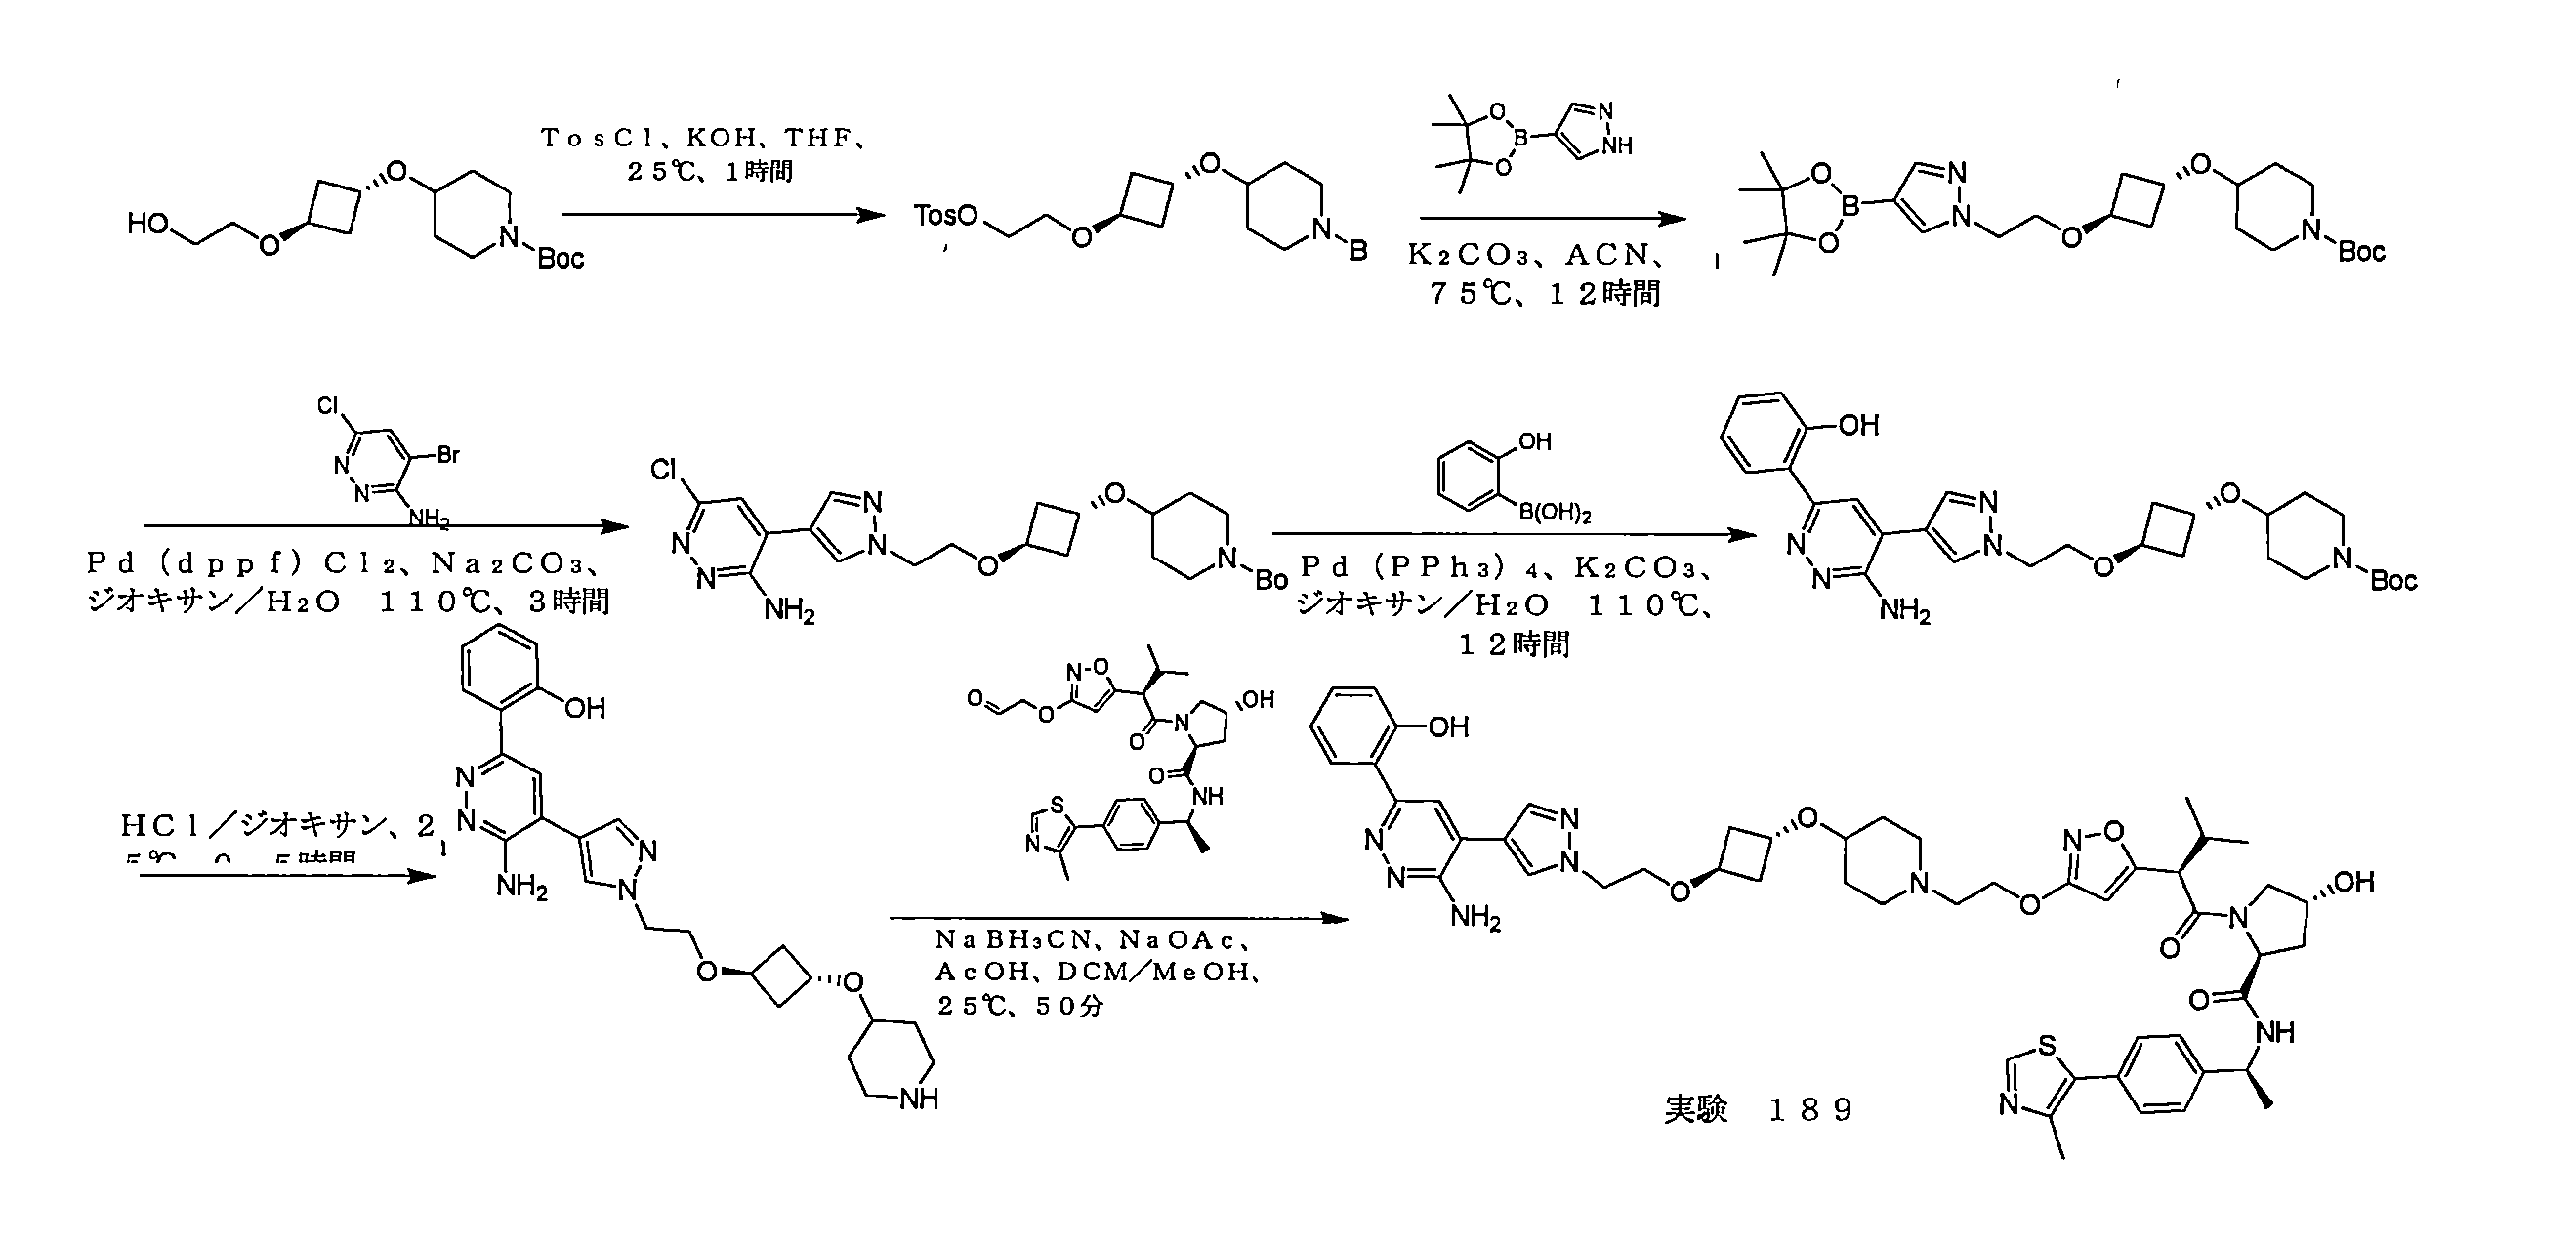
例示的な化合物194の例示的な合成

工程1

テトラヒドロフラン(50mL)中の1,4-ジメチル-1H-ピラゾール(2.5g、26.0mmol、1当量)の溶液に、ヘキサン溶液(ヘキサン中2.5M、78.0mmol、31mL、3当量)中のn-ブチルリチウムを窒素雰囲気下で、0℃で、滴下で添加した。得られた溶液を、室温で1時間撹拌し、次いで-78℃に冷却した。2-イソプロポキシ-4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン(14.52g、78.017mmol、3当量)を添加し、反応混合物を-78℃で0.5時間撹拌し、次いで0℃までゆっくり温めた。反応物をブラインでクエンチし、EtOAc(250mL×3)で抽出した。合わせた有機層をブラインで洗浄し、Na2SO4で乾燥させ、減圧下で濃縮した。残渣を、酢酸エチル/石油エーテル(1:5)で溶出するシリカゲルカラム上に適用した。これにより、2.6g(45%)の1,4-ジメチル-5-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)-1H-ピラゾールを白色の固体として得た。 To a solution of 1,4-dimethyl-1H-pyrazole (2.5 g, 26.0 mmol, 1 eq.) in tetrahydrofuran (50 mL), n-butyllithium in hexane solution (2.5 M in hexane, 78.0 mmol, 31 mL, 3 eq.) was added dropwise at 0°C under nitrogen atmosphere. The resulting solution was stirred at room temperature for 1 h and then cooled to -78°C. 2-Isopropoxy-4,4,5,5-tetramethyl-1,3,2-dioxaborolane (14.52 g, 78.017 mmol, 3 eq.) was added and the reaction mixture was stirred at -78°C for 0.5 h and then slowly warmed to 0°C. The reaction was quenched with brine and extracted with EtOAc (250 mL x 3). The combined organic layers were washed with brine, dried over Na2SO4 and concentrated under reduced pressure. The residue was applied onto a silica gel column eluting with ethyl acetate/petroleum ether (1:5). This yielded 2.6 g (45%) of 1,4-dimethyl-5-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)-1H-pyrazole as a white solid.
工程2

250mLの丸底フラスコに、ジオキサン(16mL)およびH2O(2mL)中のtert-ブチルN-[(1S)-1-(4-ブロモフェニル)エチル]カルバメート(1.0g、3.331mmol、1当量)、1,4-ジメチル-5-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)-1H-ピラゾール(1.48g、6.664mmol、2.00当量)、K2CO3(1.38g、9.963mmol、2.99当量)、およびPd(dppf)Cl2(244.0mg、0.333mmol、0.10当量)を入れた。得られた混合物を窒素雰囲気下で、90℃で3時間撹拌した。混合物を減圧下で濃縮した。残渣を、酢酸エチル/石油エーテル(1:1)で溶出するシリカゲルカラム上に適用した。これにより、1.63g(99%)のベンジルN-[(1S)-1-(4-ブロモフェニル)エチル]カルバメートを淡い黄色の固体として得た。 A 250 mL round-bottom flask was charged with tert-butyl N-[(1S)-1-(4-bromophenyl)ethyl]carbamate (1.0 g, 3.331 mmol, 1 equiv.), 1,4-dimethyl-5-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)-1H-pyrazole (1.48 g, 6.664 mmol, 2.00 equiv.), K2CO3 (1.38 g, 9.963 mmol, 2.99 equiv.), and Pd(dppf)Cl2 (244.0 mg, 0.333 mmol, 0.10 equiv.) in dioxane (16 mL) and H2O (2 mL). The resulting mixture was stirred at 90 °C for 3 h under nitrogen atmosphere. The mixture was concentrated under reduced pressure. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (1:1). This yielded 1.63 g (99%) of benzyl N-[(1S)-1-(4-bromophenyl)ethyl]carbamate as a pale yellow solid.
他の実施例について上に記載される手順を使用して、ベンジルN-[(1S)-1-(4-ブロモフェニル)エチル]カルバメートを最終化合物に変換した。 Benzyl N-[(1S)-1-(4-bromophenyl)ethyl]carbamate was converted to the final compound using the procedures described above for other examples.
例示的な化合物193を、類似の手順を使用して調製した。 Exemplary compound 193 was prepared using a similar procedure.
例示的な化合物144の例示的な合成

工程1
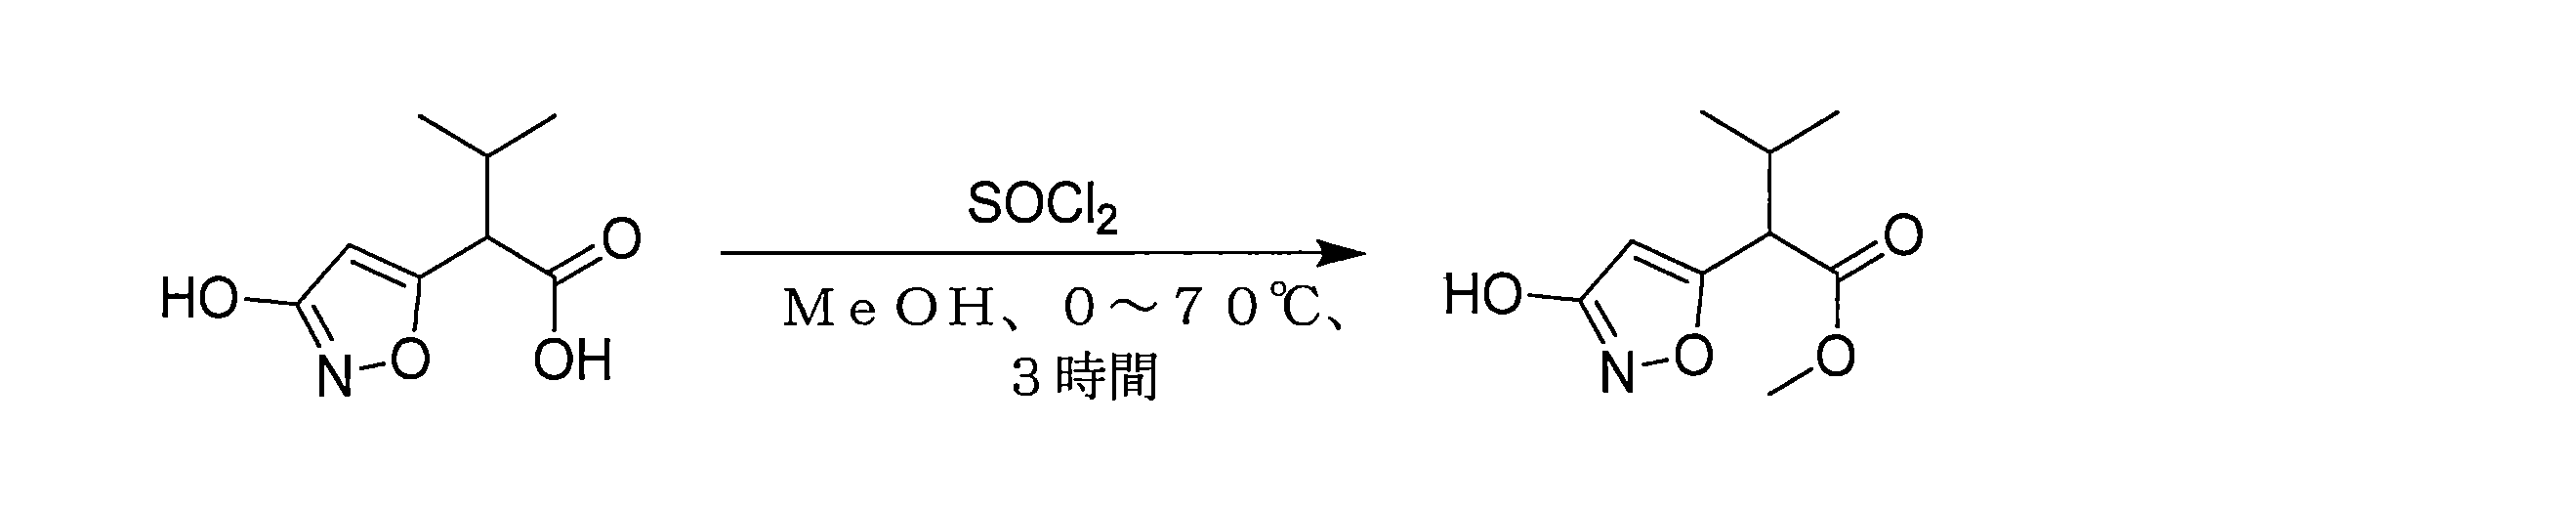
2-(3-ヒドロキシイソキサゾール-5-イル)-3-メチル-ブタン酸(1g、5.40mmol、1当量)のメタノール(10mL)溶液に、塩化チオニル(2.57g、21mmol、1.57mL、4当量)を0℃で加えた。反応混合物を3時間、70℃で撹拌した。反応混合物を減圧下で濃縮した。残渣を水(50mL)で希釈し、酢酸エチル(30mL×3)で抽出した。合わせた有機層をブライン(80mL×2)で洗浄し、無水硫酸ナトリウム上で乾燥させ、濾過し、減圧下で濃縮した。メチル2-(3-ヒドロキシイソキサゾール-5-イル)-3-メチル-ブタノエート(1g、5.02mmol、収率92%)を黄色の油として得た。 To a solution of 2-(3-hydroxyisoxazol-5-yl)-3-methyl-butanoic acid (1 g, 5.40 mmol, 1 equiv.) in methanol (10 mL) was added thionyl chloride (2.57 g, 21 mmol, 1.57 mL, 4 equiv.) at 0 °C. The reaction mixture was stirred at 70 °C for 3 h. The reaction mixture was concentrated under reduced pressure. The residue was diluted with water (50 mL) and extracted with ethyl acetate (30 mL x 3). The combined organic layers were washed with brine (80 mL x 2), dried over anhydrous sodium sulfate, filtered and concentrated under reduced pressure. Methyl 2-(3-hydroxyisoxazol-5-yl)-3-methyl-butanoate (1 g, 5.02 mmol, 92% yield) was obtained as a yellow oil.
工程2

メチル2-(3-ヒドロキシイソキサゾール-5-イル)-3-メチル-ブタノエート(800mg、4.02mmol、1当量)のアセトニトリル(5mL)溶液に、炭酸カリウム(1.11g、8.03mmol、2当量)およびペルフルオロブチルスルホニルフルオリド(1.46g、4.82mmol、1.2当量)を加えた。反応混合物を25℃で12時間撹拌した。反応混合物を水(50mL)で希釈し、酢酸エチル(30mL×3)で抽出した。合わせた有機層をブライン(80mL×3)で洗浄し、無水硫酸ナトリウム上で乾燥させ、濾過し、減圧下で濃縮した。残渣を、シリカゲルカラムクロマトグラフィー(石油エーテル/酢酸エチル=100/1~20/1)によって精製した。メチル3-メチル-2-[3-(1,1,2,2,3,3,4,4,4-ノナフルオロブチルスルホニルオキシ)イソキサゾール-5-イル]ブタノエート(530mg、1.10mmol、収率27%)を無色の油として得た。 To a solution of methyl 2-(3-hydroxyisoxazol-5-yl)-3-methyl-butanoate (800 mg, 4.02 mmol, 1 eq.) in acetonitrile (5 mL), potassium carbonate (1.11 g, 8.03 mmol, 2 eq.) and perfluorobutylsulfonyl fluoride (1.46 g, 4.82 mmol, 1.2 eq.) were added. The reaction mixture was stirred at 25°C for 12 h. The reaction mixture was diluted with water (50 mL) and extracted with ethyl acetate (30 mL x 3). The combined organic layers were washed with brine (80 mL x 3), dried over anhydrous sodium sulfate, filtered and concentrated under reduced pressure. The residue was purified by silica gel column chromatography (petroleum ether/ethyl acetate = 100/1 to 20/1). Methyl 3-methyl-2-[3-(1,1,2,2,3,3,4,4,4-nonafluorobutylsulfonyloxy)isoxazol-5-yl]butanoate (530 mg, 1.10 mmol, 27% yield) was obtained as a colorless oil.
工程3

メチル3-メチル-2-[3-(1,1,2,2,3,3,4,4,4-ノナフルオロブチルスルホニルオキシ)イソキサゾール-5-イル]ブタノエート(400mg、0.83mmol、1当量)およびtert-ブチルピペラジン-1-カルボキシレート(154mg、0.83mmol、1当量)のジメチルホルムアミド(5mL)溶液を、130℃で12時間撹拌した。反応混合物を、水(30mL)で希釈し、酢酸エチル(30mL×3)で抽出した。合わせた有機層をブライン(30mL×3)で洗浄し、無水硫酸ナトリウム上で乾燥させ、濾過し、減圧下で濃縮した。残渣を分取TLC(石油エーテル:酢酸エチル=3:1)によって精製した。Tert-ブチル4-[5-(1-メトキシカルボニル-2-メチル-プロピル)イソキサゾール-3-イル]ピペラジン-1-カルボキシレート(130mg、0.35mmol、収率42%)を無色の油として得た。 A solution of methyl 3-methyl-2-[3-(1,1,2,2,3,3,4,4,4-nonafluorobutylsulfonyloxy)isoxazol-5-yl]butanoate (400 mg, 0.83 mmol, 1 eq.) and tert-butyl piperazine-1-carboxylate (154 mg, 0.83 mmol, 1 eq.) in dimethylformamide (5 mL) was stirred at 130 °C for 12 h. The reaction mixture was diluted with water (30 mL) and extracted with ethyl acetate (30 mL x 3). The combined organic layer was washed with brine (30 mL x 3), dried over anhydrous sodium sulfate, filtered, and concentrated under reduced pressure. The residue was purified by preparative TLC (petroleum ether: ethyl acetate = 3:1). Tert-butyl 4-[5-(1-methoxycarbonyl-2-methyl-propyl)isoxazol-3-yl]piperazine-1-carboxylate (130 mg, 0.35 mmol, 42% yield) was obtained as a colorless oil.
工程4

メタノール(2mL)、テトラヒドロフラン(2mL)、および水(2mL)中のtert-ブチル4-[5-(1-メトキシカルボニル-2-メチル-プロピル)イソキサゾール-3-イル]ピペラジン-1-カルボキシレート(130mg、0.35mmol、1当量)の溶液に、水酸化リチウム(44mg、1.06mmol、3当量)を添加した。反応混合物を、15℃で1時間撹拌した。水(3mL)を加えた。pHを塩酸(水中1M)で6に調整し、酢酸エチル(10mL×5)で抽出した。合わせた有機層を、ブライン(50mL×1)で洗浄し、無水硫酸ナトリウム上で乾燥させ、濾過し、減圧下で濃縮した。2-[3-(4-tert-ブトキシカルボニルピペラジン-1-イル)イソキサゾール-5-イル]-3-メチル-ブタン酸(100mg、0.28mmol、収率79%)を白色の固体として得た。 To a solution of tert-butyl 4-[5-(1-methoxycarbonyl-2-methyl-propyl)isoxazol-3-yl]piperazine-1-carboxylate (130 mg, 0.35 mmol, 1 equiv.) in methanol (2 mL), tetrahydrofuran (2 mL), and water (2 mL) was added lithium hydroxide (44 mg, 1.06 mmol, 3 equiv.). The reaction mixture was stirred at 15°C for 1 h. Water (3 mL) was added. The pH was adjusted to 6 with hydrochloric acid (1M in water) and extracted with ethyl acetate (10 mL x 5). The combined organic layers were washed with brine (50 mL x 1), dried over anhydrous sodium sulfate, filtered, and concentrated under reduced pressure. 2-[3-(4-tert-butoxycarbonylpiperazin-1-yl)isoxazol-5-yl]-3-methyl-butanoic acid (100 mg, 0.28 mmol, 79% yield) was obtained as a white solid.
工程5

ジメチルホルムアミド(5mL)中の2-[3-(4-tert-ブトキシカルボニルピペラジン-1-イル)イソキサゾール-5-イル]-3-メチル-ブタン酸(100mg、0.28mmol、1当量)およびO-(7-アザベンゾトリアゾール-1-イル)-N,N,N’,N’-テトラメチルウロニウムヘキサフルオロホスフェート(129mg、0.33mmol、1.2当量)の溶液に、(2S,4R)-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド塩酸塩(104mg、0.28mmol、1当量)およびジイソプロピルエチルアミン(109mg、0.84mmol、3当量)を添加した。反応混合物を15℃で0.5時間撹拌した。反応混合物を水(20mL)の中へと注ぎ、酢酸エチル(30mL×3)で抽出した。合わせた有機層をブライン(50mL×3)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、減圧下で濃縮した。残渣を、分取TLC(ジクロロメタン:メタノール=10:1)によって精製した。tert-ブチル4-[5-[1-[(2S,4R)-4-ヒドロキシ-2-[[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]カルバモイル]ピロリジン-1-カルボニル]-2-メチル-プロピル]イソキサゾール-3-イル]ピペラジン-1-カルボキシレート(180mg、0.26mmol、収率92%)を無色の油として得た。 To a solution of 2-[3-(4-tert-butoxycarbonylpiperazin-1-yl)isoxazol-5-yl]-3-methyl-butanoic acid (100 mg, 0.28 mmol, 1 equiv.) and O-(7-azabenzotriazol-1-yl)-N,N,N',N'-tetramethyluronium hexafluorophosphate (129 mg, 0.33 mmol, 1.2 equiv.) in dimethylformamide (5 mL), (2S,4R)-4-hydroxy-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide hydrochloride (104 mg, 0.28 mmol, 1 equiv.) and diisopropylethylamine (109 mg, 0.84 mmol, 3 equiv.) were added. The reaction mixture was stirred at 15°C for 0.5 h. The reaction mixture was poured into water (20 mL) and extracted with ethyl acetate (30 mL x 3). The combined organic layers were washed with brine (50 mL x 3), dried over anhydrous sodium sulfate, filtered, and concentrated under reduced pressure. The residue was purified by preparative TLC (dichloromethane:methanol = 10:1). tert-Butyl 4-[5-[1-[(2S,4R)-4-hydroxy-2-[[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]carbamoyl]pyrrolidine-1-carbonyl]-2-methyl-propyl]isoxazol-3-yl]piperazine-1-carboxylate (180 mg, 0.26 mmol, 92% yield) was obtained as a colorless oil.
工程6
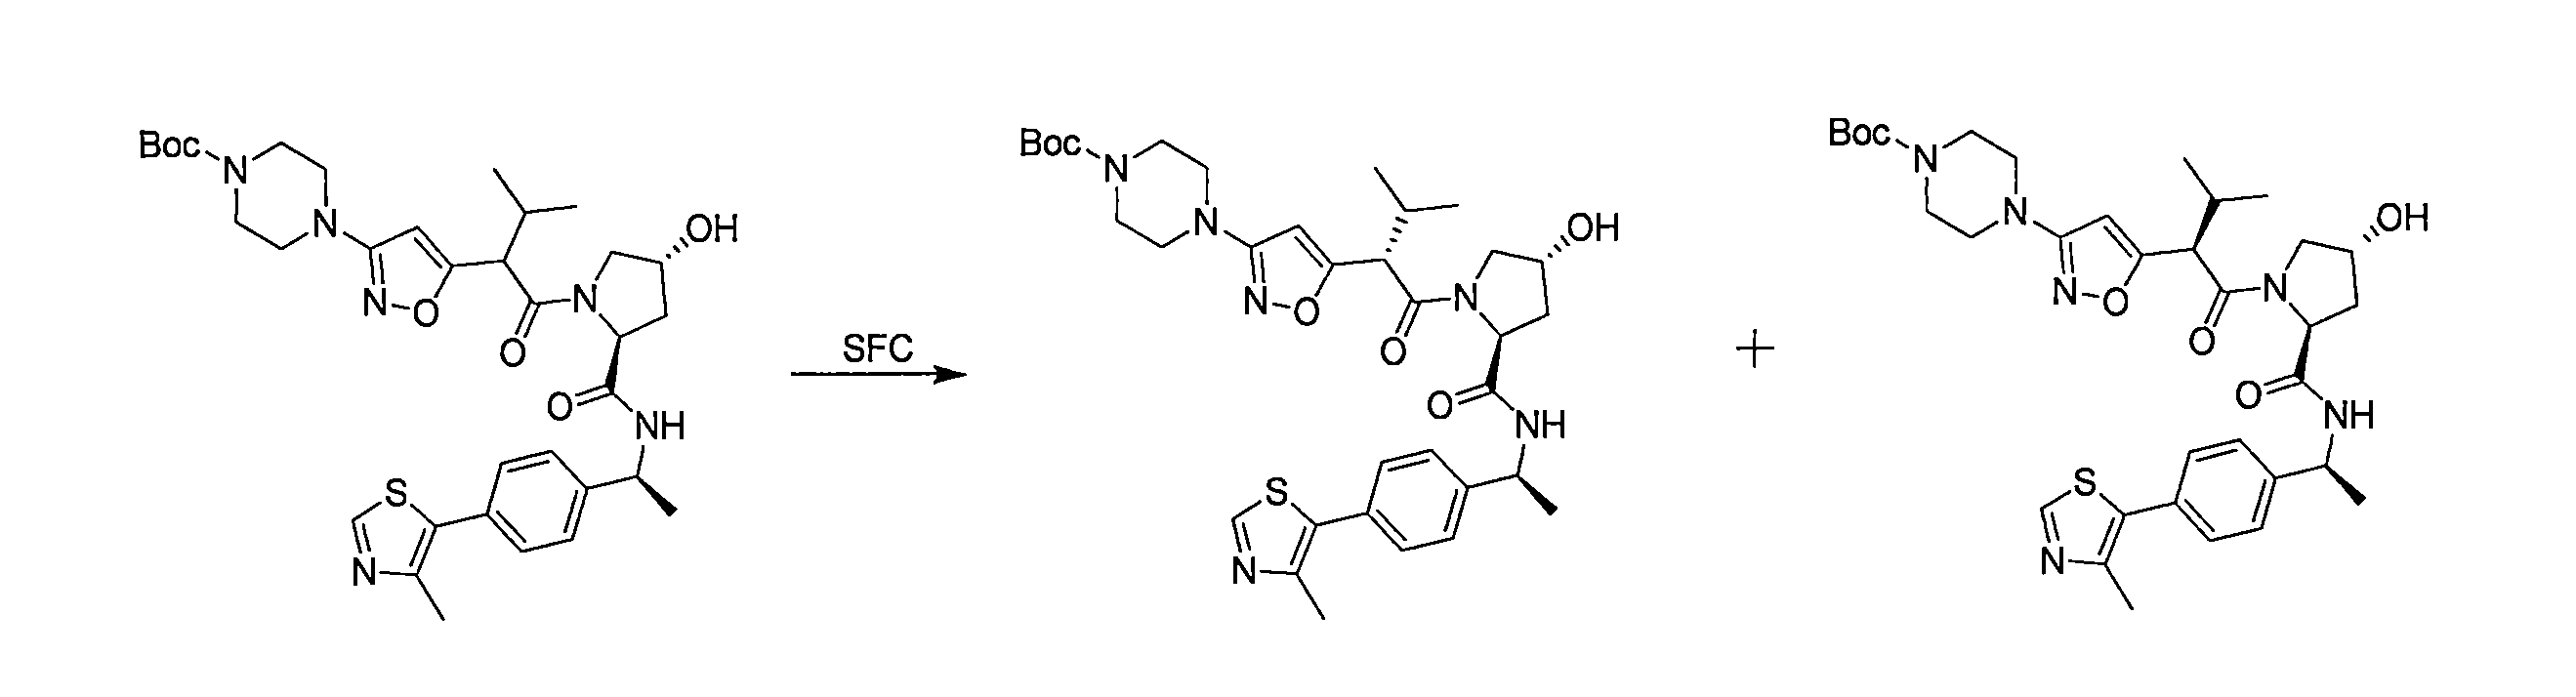
tert-ブチル4-[5-[1-[(2S,4R)-4-ヒドロキシ-2-[[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]カルバモイル]ピロリジン-1-カルボニル]-2-メチル-プロピル]イソキサゾール-3-イル]ピペラジン-1-カルボキシレート(200mg、0.29mmol、1当量)を、キラル超臨界流体クロマトグラフィーによって分離した。機器:CASWH-Prep-SFC-C、SFC Column:DAICEL Chiralpak AS(250mm*30mm、10um);移動相:CO2中のエタノール(0.1% NH3.H2O)、25%~25%;流速:55g/分、波長:220nm。tert-ブチル4-[5-[(1S)-1-[(2S,4R)-4-ヒドロキシ-2-[[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]カルバモイル]ピロリジン-1-カルボニル]-2-メチル-プロピル]イソキサゾール-3-イル]ピペラジン-1-カルボキシレート(80mg、0.11mmol、収率76%、純度95%)を白色の固体として得た。tert-ブチル4-[5-[(1R)-1-[(2S,4R)-4-ヒドロキシ-2-[[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]カルバモイル]ピロリジン-1-カルボニル]-2-メチル-プロピル]イソキサゾール-3-イル]ピペラジン-1-カルボキシレート(70mg、0.1mmol、収率68%、純度98%)を白色の固体として得た。 tert-Butyl 4-[5-[1-[(2S,4R)-4-hydroxy-2-[[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]carbamoyl]pyrrolidine-1-carbonyl]-2-methyl-propyl]isoxazol-3-yl]piperazine-1-carboxylate (200 mg, 0.29 mmol, 1 equiv.) was separated by chiral supercritical fluid chromatography. Instrument: CASWH-Prep-SFC-C, SFC Column: DAICEL Chiralpak AS (250 mm*30 mm, 10 um); Mobile phase: Ethanol in CO2 (0.1% NH3.H2O), 25%-25%; Flow rate: 55 g/min, Wavelength: 220 nm. Obtained tert-butyl 4-[5-[(1S)-1-[(2S,4R)-4-hydroxy-2-[[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]carbamoyl]pyrrolidine-1-carbonyl]-2-methyl-propyl]isoxazol-3-yl]piperazine-1-carboxylate (80 mg, 0.11 mmol, 76% yield, 95% purity) as a white solid. tert-Butyl 4-[5-[(1R)-1-[(2S,4R)-4-hydroxy-2-[[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]carbamoyl]pyrrolidine-1-carbonyl]-2-methyl-propyl]isoxazol-3-yl]piperazine-1-carboxylate (70 mg, 0.1 mmol, 68% yield, 98% purity) was obtained as a white solid.
工程7
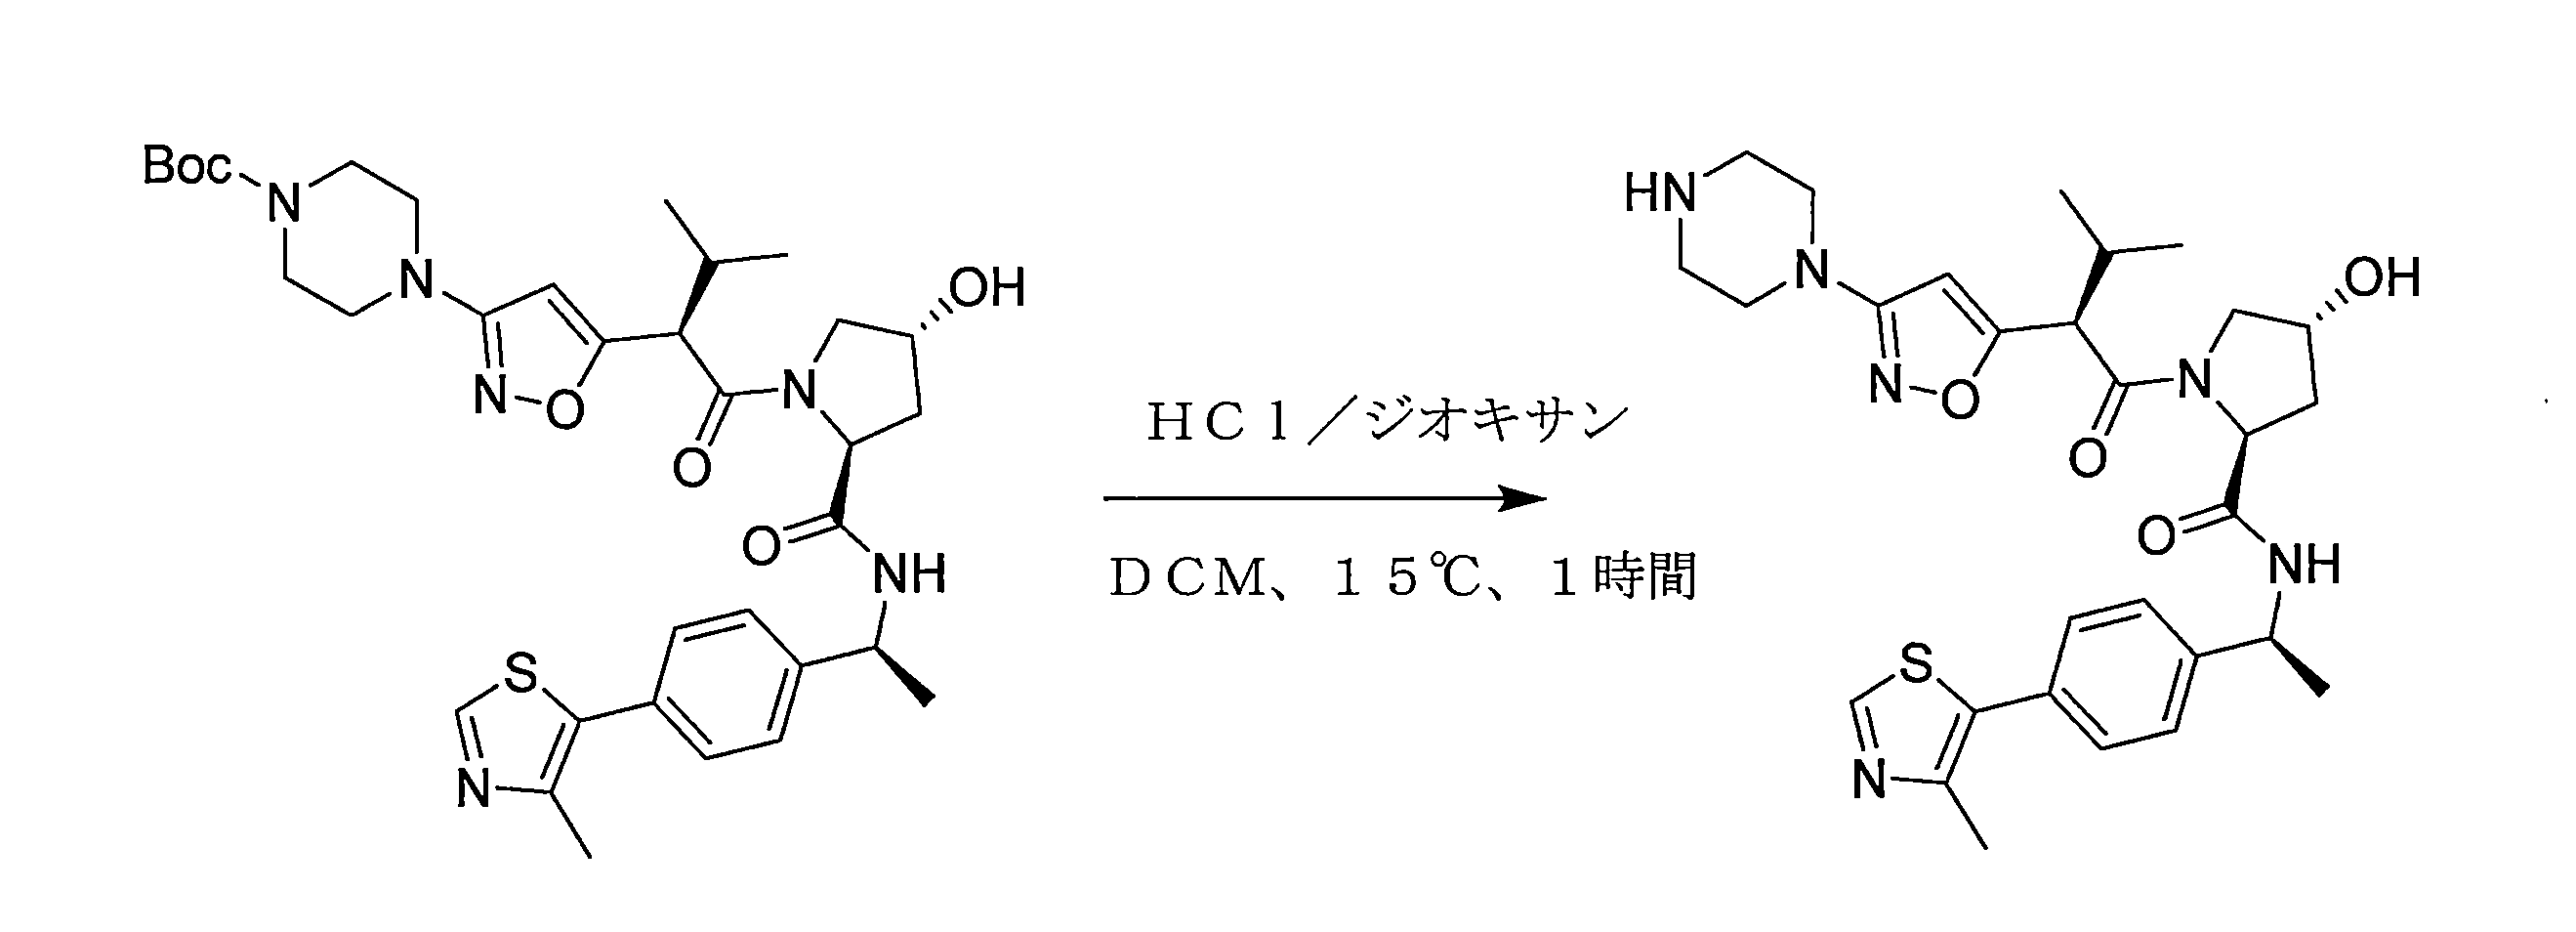
ジクロロメタン(2mL)中のtert-ブチル4-[5-[(1R)-1-[(2S,4R)-4-ヒドロキシ-2-[[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]カルバモイル]ピロリジン-1-カルボニル]-2-メチル-プロピル]イソキサゾール-3-イル]ピペラジン-1-カルボキシレート(70mg、0.1mmol、1当量)の溶液に、塩酸(ジオキサン中4M、3mL)を加えた。反応混合物を15℃で1時間撹拌した。反応混合物を減圧下で濃縮した。(2S,4R)-4-ヒドロキシ-1-[(2R)-3-メチル-2-(3-ピペラジン-1-イルイソキサゾール-5-イル)ブタノイル]-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(55mg、0.09mmol、収率91%)を白色の固体として得た。 To a solution of tert-butyl 4-[5-[(1R)-1-[(2S,4R)-4-hydroxy-2-[[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]carbamoyl]pyrrolidine-1-carbonyl]-2-methyl-propyl]isoxazol-3-yl]piperazine-1-carboxylate (70 mg, 0.1 mmol, 1 equiv.) in dichloromethane (2 mL) was added hydrochloric acid (4 M in dioxane, 3 mL). The reaction mixture was stirred at 15°C for 1 h. The reaction mixture was concentrated under reduced pressure. (2S,4R)-4-hydroxy-1-[(2R)-3-methyl-2-(3-piperazin-1-ylisoxazol-5-yl)butanoyl]-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (55 mg, 0.09 mmol, 91% yield) was obtained as a white solid.
工程8
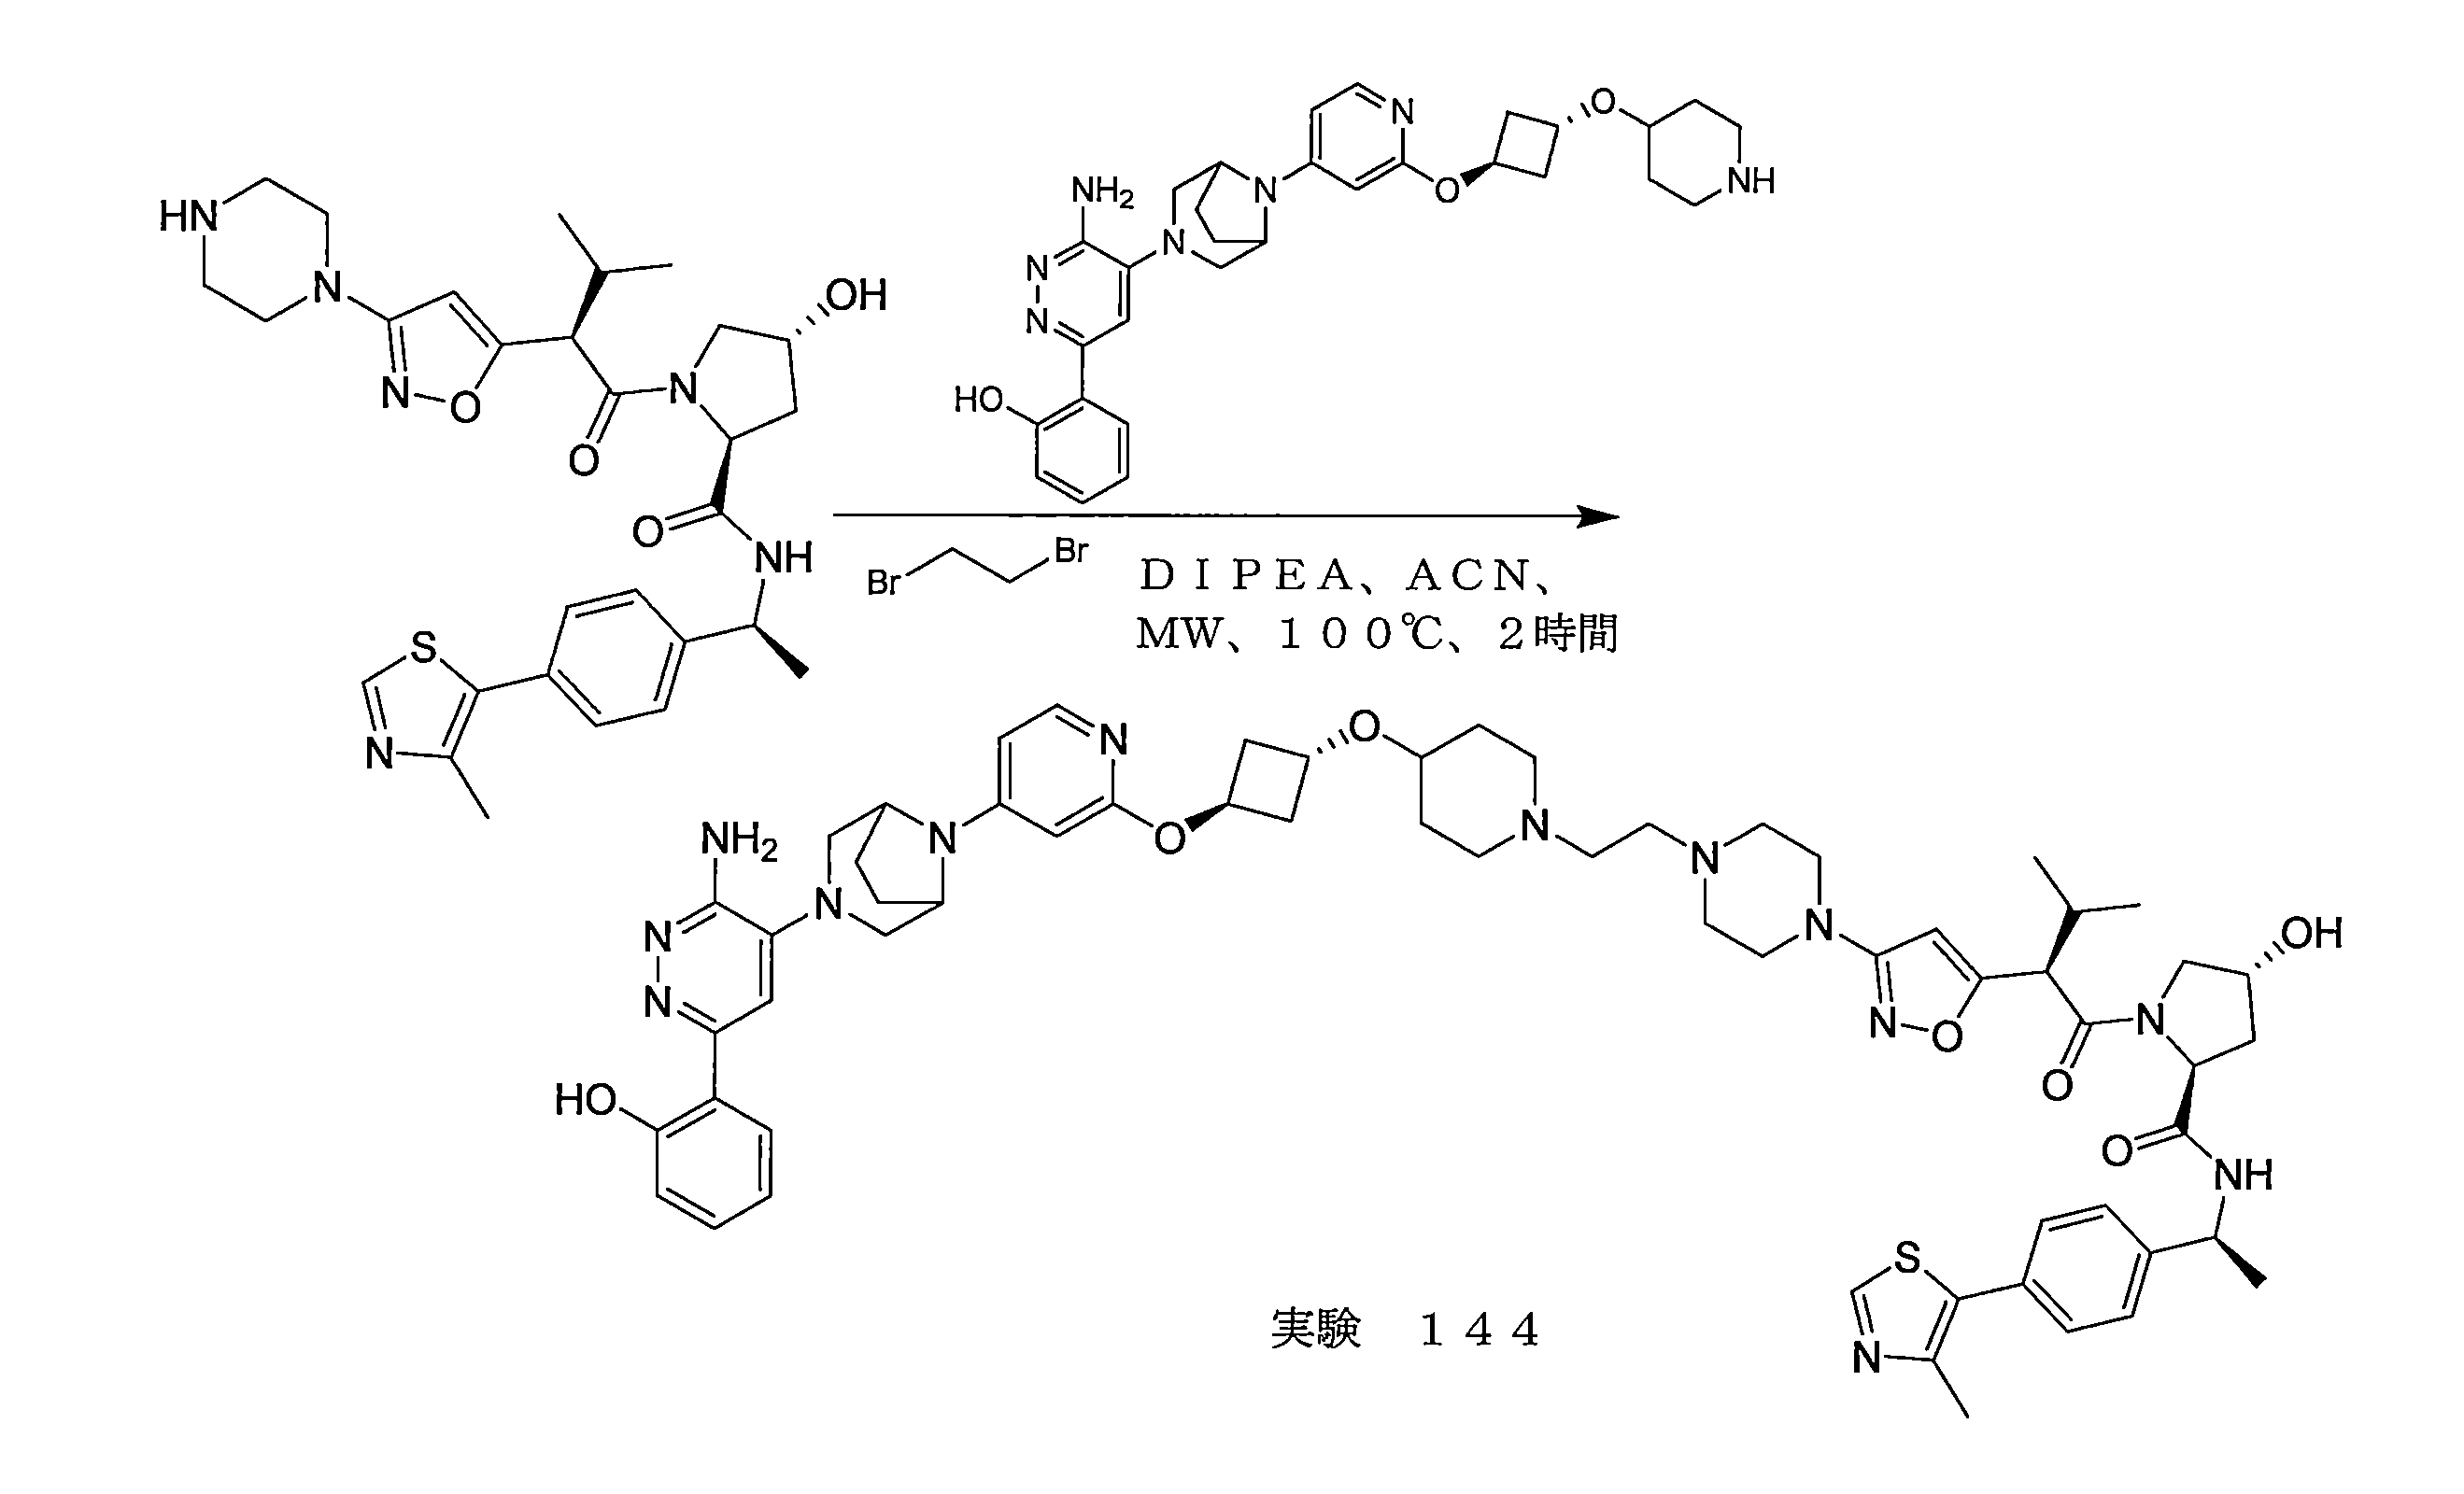
アセトニトリル(12mL)中の(2S,4R)-4-ヒドロキシ-1-[(2R)-3-メチル-2-(3-ピペラジン-1-イルイソキサゾール-5-イル)ブタノイル]-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(100mg、0.16mmol、1当量、塩酸塩)、1,2-ジブロモエタン(31mg、0.16mmol、1当量)、および2-[6-アミノ-5-[8-[2-[3-(4-ピペリジルオキシ)シクロブトキシ]-4-ピリジル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-イル]ピリダジン-3-イル]フェノール二塩酸塩(102mg、0.16mmol、1当量)の混合物に、N,N-ジイソプロピルエチルアミン(128mg、0.99mmol、6当量)を添加した。混合物をマイクロ波下で、100℃で2時間撹拌した。反応混合物を減圧下で濃縮して残渣を得た。残渣を、分取HPLC(カラム:Phenomenex Synergi C18 150*25*10um;移動相:[水(0.225%FA)-ACN];B%:5%~35%、10分)によって精製した。化合物(2S,4R)-1-[(2R)-2-[3-[4-[2-[4-[3-[[4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]-2-ピリジル]オキシ]シクロブトキシ]-1-ピペリジル]エチル]ピペラジン-1-イル]イソキサゾール-5-イル]-3-メチル-ブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミドギ酸塩(24.6mg、0.02mmol、収率12%)を、灰色がかった白色の固体として得た。 (2S,4R)-4-hydroxy-1-[(2R)-3-methyl-2-(3-piperazin-1-ylisoxazol-5-yl)butanoyl]-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (100 mg, 0.16 mmol, 1 equiv., hydrochloride salt), 1,2-dibromoethane (31 mg, 0.1 To a mixture of 2-[6-amino-5-[8-[2-[3-(4-piperidyloxy)cyclobutoxy]-4-pyridyl]-3,8-diazabicyclo[3.2.1]octan-3-yl]pyridazin-3-yl]phenol dihydrochloride (102 mg, 0.16 mmol, 1 eq.), N,N-diisopropylethylamine (128 mg, 0.99 mmol, 6 eq.) was added. The mixture was stirred at 100° C. under microwave for 2 hours. The reaction mixture was concentrated under reduced pressure to give a residue. The residue was purified by preparative HPLC (Column: Phenomenex Synergi C18 150*25*10 um; Mobile phase: [Water (0.225% FA)-ACN]; B%: 5%-35%, 10 min). The compound (2S,4R)-1-[(2R)-2-[3-[4-[2-[4-[3-[[4-[3-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl]-2-pyridyl]oxy]cyclobutoxy]-1-piperidyl]ethyl]piperazin-1-yl]isoxazol-5-yl]-3-methyl-butanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide formate (24.6 mg, 0.02 mmol, 12% yield) was obtained as an off-white solid.
例示的な化合物195として類似の手順を使用して調製した。 Prepared using a similar procedure as exemplary compound 195.
例示的な化合物197および352の例示的な合成
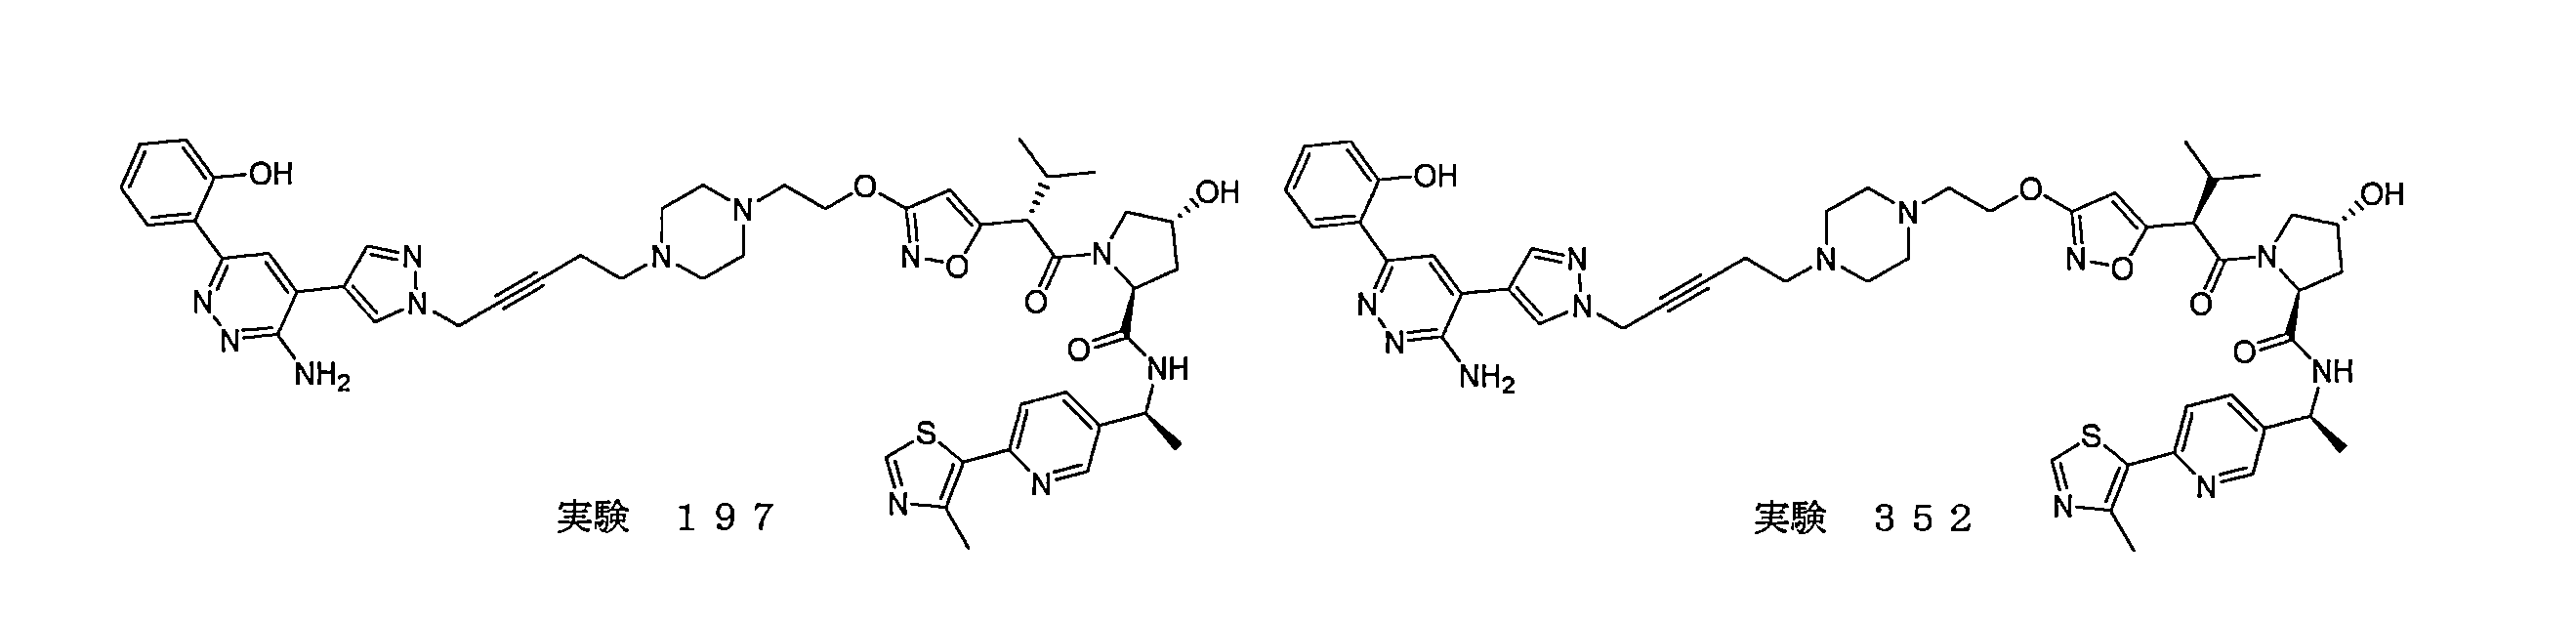
例示的な化合物112および116について記載されるものと類似の手順を使用して調製した。 Prepared using procedures similar to those described for exemplary compounds 112 and 116.
例示的な化合物114の例示的な合成
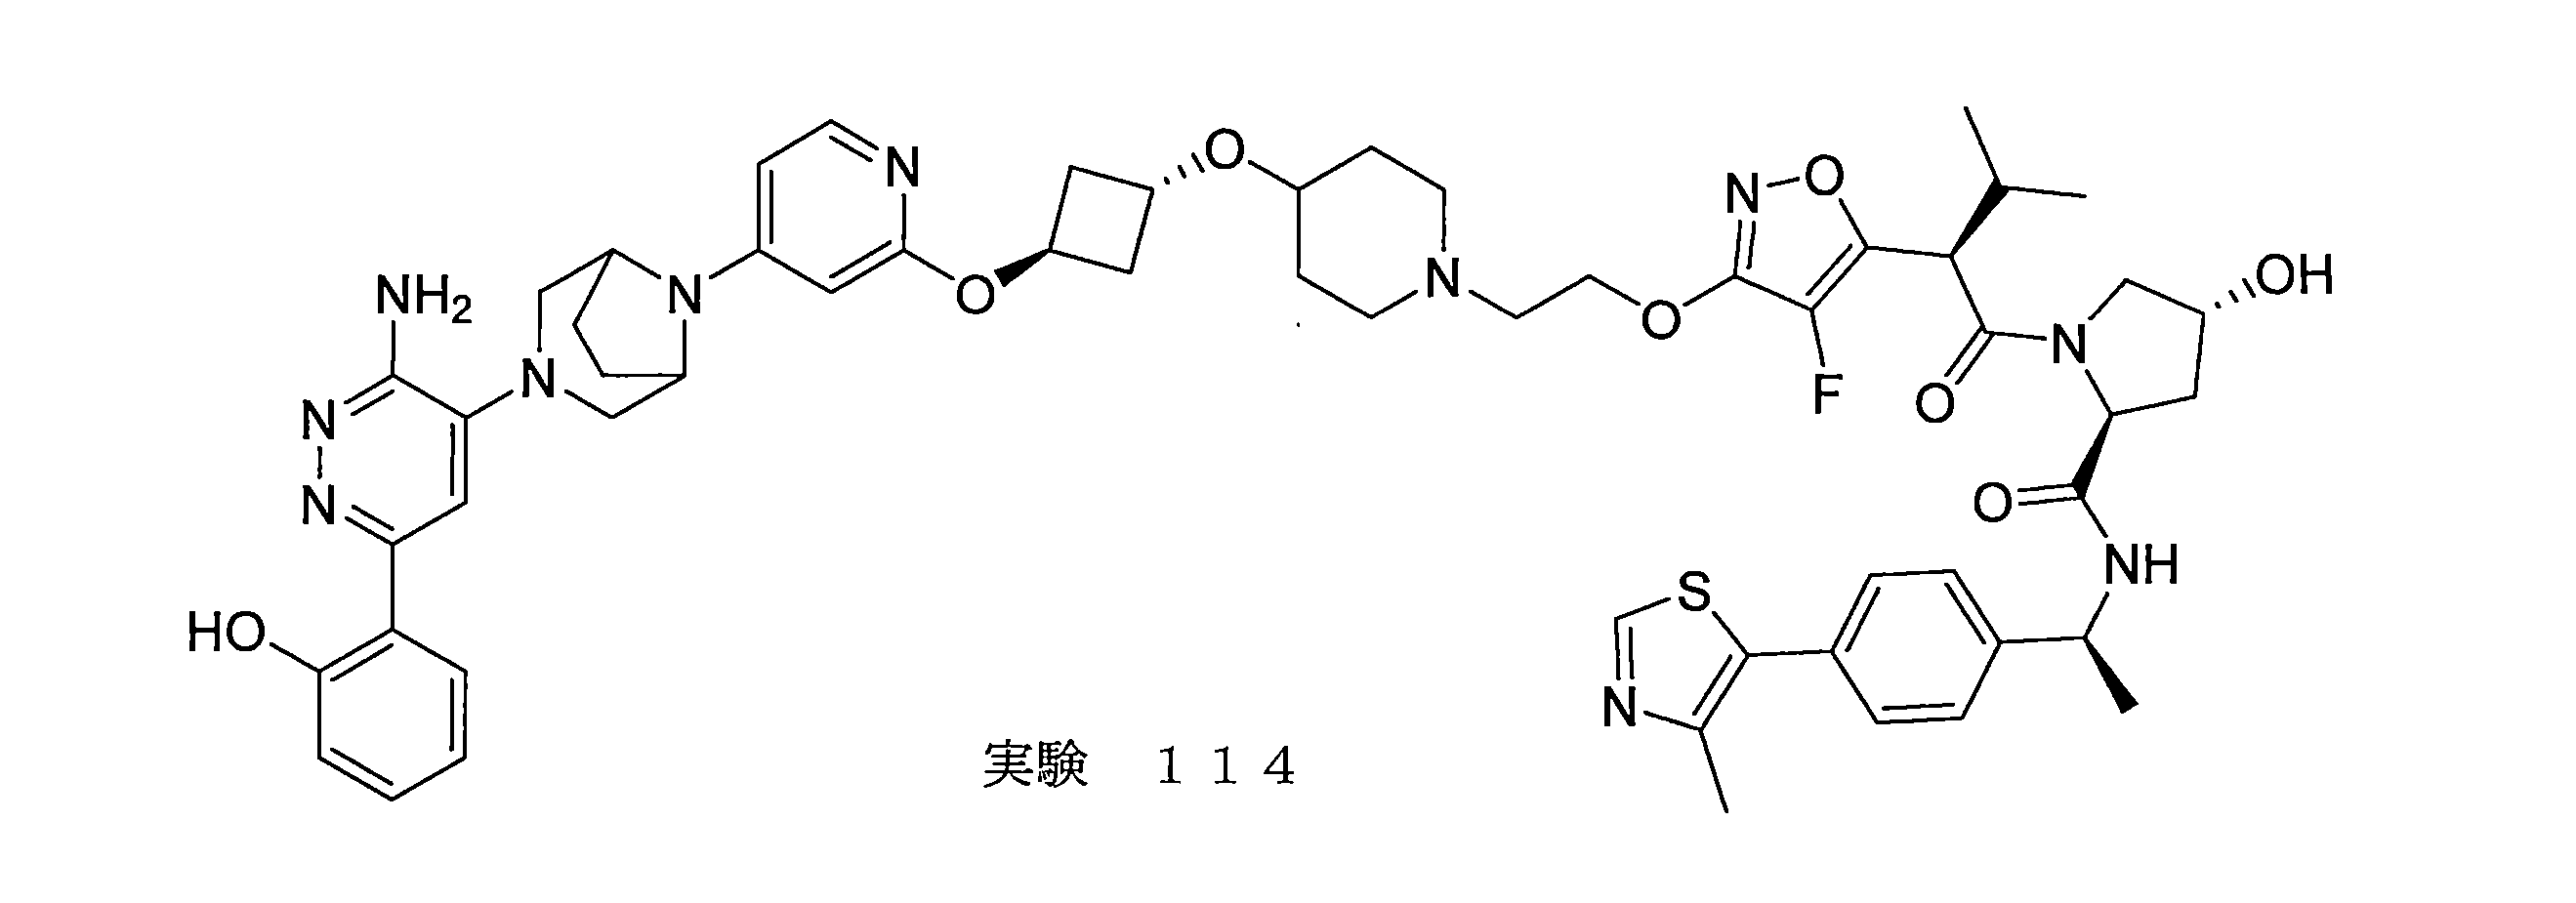
工程1

トルエン(500mL)中のエチル2-フルオロ-3-オキソ-ブタノエート(33.6g、226.82mmol、28.47mL、1当量)およびエチレングリコール(70.39g、1.13mol、63.42mL、5当量)の溶液に、三フッ化ホウ素エチルエーテル(3.36g、23.67mmol、2.92mL、0.1当量)を添加した。混合物を80℃で12時間撹拌した。反応混合物を水(300mL)で希釈し、酢酸エチル(300mL)で2回抽出した。合わせた有機相をブライン(300mL)で2回洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、濃縮して、粗生成物を得た。残渣を、シリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=100:1~10:1)により精製した。化合物エチル2-フルオロ-2-(2-メチル-1,3-ジオキソラン-2-イル)酢酸塩(29.3g、152.5mmol、収率67%)を無色の油として得た。 To a solution of ethyl 2-fluoro-3-oxo-butanoate (33.6 g, 226.82 mmol, 28.47 mL, 1 eq.) and ethylene glycol (70.39 g, 1.13 mol, 63.42 mL, 5 eq.) in toluene (500 mL), boron trifluoride ethyl etherate (3.36 g, 23.67 mmol, 2.92 mL, 0.1 eq.) was added. The mixture was stirred at 80 °C for 12 h. The reaction mixture was diluted with water (300 mL) and extracted twice with ethyl acetate (300 mL). The combined organic phase was washed twice with brine (300 mL), dried over anhydrous sodium sulfate, filtered, and concentrated to give the crude product. The residue was purified by silica gel chromatography (petroleum ether: ethyl acetate = 100: 1 to 10: 1). The compound ethyl 2-fluoro-2-(2-methyl-1,3-dioxolan-2-yl)acetate (29.3 g, 152.5 mmol, yield 67%) was obtained as a colorless oil.
工程2

ピリジン(164mL)中のエチル2-フルオロ-2-(2-メチル-1,3-ジオキソラン-2-イル)酢酸塩(29.3g、152.46mmol、1当量)およびヒドロキシルアミン塩酸塩(21.19g、304.92mmol、2当量)の溶液に、メタノール(61mL)中のメトキシドナトリウム(28%)を1時間にわたって0℃で、滴下で添加した。混合物を20℃で1時間撹拌した。反応混合物を濾過し、濾液をテトラヒドロフラン(500mL)で希釈し、さらに濾過した。濾液を濃縮して、粗生成物を得、これをシリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=1:0~0:1)によって精製して、生成物を得た。化合物2-フルオロ-2-(2-メチル-1,3-ジオキソラン-2-イル)エタンヒドロキサム酸(10.4g、58.1mmol、収率38%)をオレンジ色の油として得た。 To a solution of ethyl 2-fluoro-2-(2-methyl-1,3-dioxolan-2-yl)acetate (29.3 g, 152.46 mmol, 1 eq.) and hydroxylamine hydrochloride (21.19 g, 304.92 mmol, 2 eq.) in pyridine (164 mL), sodium methoxide (28%) in methanol (61 mL) was added dropwise over 1 h at 0 °C. The mixture was stirred at 20 °C for 1 h. The reaction mixture was filtered, and the filtrate was diluted with tetrahydrofuran (500 mL) and further filtered. The filtrate was concentrated to give the crude product, which was purified by silica gel chromatography (petroleum ether:ethyl acetate = 1:0 to 0:1) to give the product. Compound 2-fluoro-2-(2-methyl-1,3-dioxolan-2-yl)ethanehydroxamic acid (10.4 g, 58.1 mmol, 38% yield) was obtained as an orange oil.
工程3

硫酸(18M、50mL、15.58当量)中の2-フルオロ-2-(2-メチル-1,3-ジオキソラン-2-イル)エタンヒドロキサム酸(10.35g、57.77mmol、1当量)の溶液を、70℃で2時間撹拌した。反応混合物を氷水(250mL)に注ぎ入れ、次いで、酢酸エチル(250mL)で2回抽出した。合わせた有機層を無水硫酸ナトリウムで乾燥させ、濃縮して、粗生成物を得た。粗生成物を、精製することなく次の工程で直接使用した。化合物4-フルオロ-5-メチル-イソキサゾール-3-オール(4.60g、39.29mmol)を黄色の固体として得た。 A solution of 2-fluoro-2-(2-methyl-1,3-dioxolan-2-yl)ethanehydroxamic acid (10.35 g, 57.77 mmol, 1 eq.) in sulfuric acid (18 M, 50 mL, 15.58 eq.) was stirred at 70 °C for 2 h. The reaction mixture was poured into ice water (250 mL) and then extracted twice with ethyl acetate (250 mL). The combined organic layers were dried over anhydrous sodium sulfate and concentrated to give the crude product. The crude product was used directly in the next step without purification. Compound 4-fluoro-5-methyl-isoxazol-3-ol (4.60 g, 39.29 mmol) was obtained as a yellow solid.
工程4

ジクロロメタン(228mL)中の4-フルオロ-5-メチル-イソキサゾール-3-オール(5.7g、48.69mmol、1当量)の溶液に、p-トルエンスルホン酸一水和物(926mg、4.87mmol、0.1当量)および3,4-ジヒドロ-2H-ピラン(20.48g、243.43mmol、22.26mL、5当量)を添加した。混合物を20℃で10時間撹拌した。反応混合物を水(100mL)に注ぎ入れ、pHを飽和重炭酸ナトリウム溶液(60mL)で7に調整した。次いで、混合物をジクロロメタン(200mL)で2回抽出した。合わせた有機層を飽和塩化ナトリウム水溶液(200mL)で2回洗浄し、無水硫酸ナトリウムで乾燥させ、濾過した。濾液を濃縮して、粗生成物を得、これをシリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=1:0~8:1)によって精製した。化合物4-フルオロ-5-メチル-3-テトラヒドロピラン-2-イルオキシ-イソキサゾール(7.94g、39.5mmol、収率81%)を黄色の固体として得た。 To a solution of 4-fluoro-5-methyl-isoxazol-3-ol (5.7 g, 48.69 mmol, 1 eq.) in dichloromethane (228 mL), p-toluenesulfonic acid monohydrate (926 mg, 4.87 mmol, 0.1 eq.) and 3,4-dihydro-2H-pyran (20.48 g, 243.43 mmol, 22.26 mL, 5 eq.) were added. The mixture was stirred at 20 °C for 10 h. The reaction mixture was poured into water (100 mL) and the pH was adjusted to 7 with saturated sodium bicarbonate solution (60 mL). The mixture was then extracted twice with dichloromethane (200 mL). The combined organic layers were washed twice with saturated aqueous sodium chloride solution (200 mL), dried over anhydrous sodium sulfate, and filtered. The filtrate was concentrated to give the crude product, which was purified by silica gel chromatography (petroleum ether: ethyl acetate = 1:0 to 8:1). The compound 4-fluoro-5-methyl-3-tetrahydropyran-2-yloxy-isoxazole (7.94 g, 39.5 mmol, yield 81%) was obtained as a yellow solid.
工程5
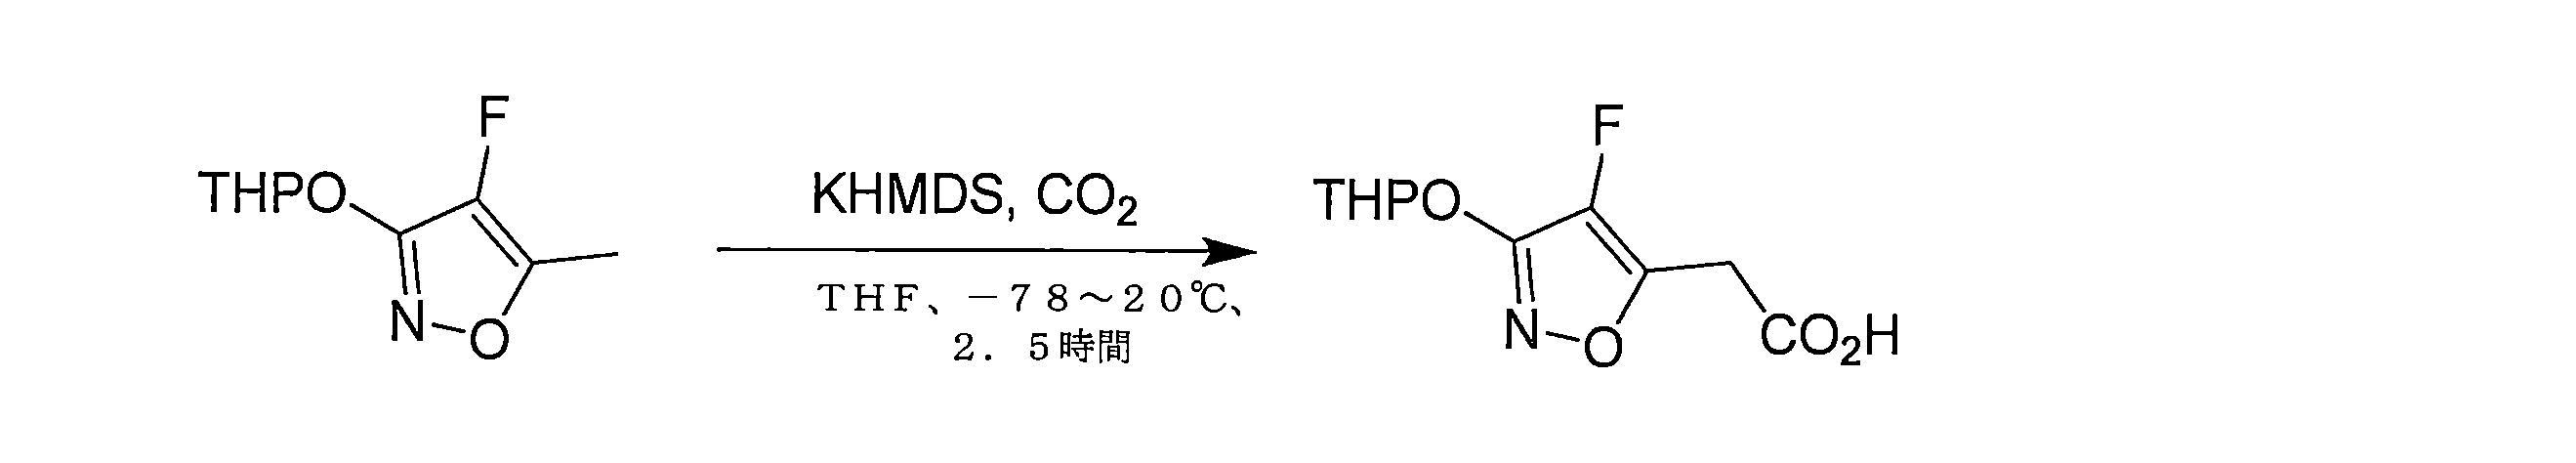
テトラヒドロフラン(115mL)中の4-フルオロ-5-メチル-3-テトラヒドロピラン-2-イルオキシ-イソキサゾール(8.07g、40.13mmol、1当量)の溶液に、カリウムビス(トリメチルシリル)アミド(1M、60.20mL、1.5当量)を窒素下で、-78℃で、滴下で添加し、反応混合物を-78℃で0.5時間撹拌した。反応混合物を通して、その後、二酸化炭素(15psi)を-78℃で1時間泡立て、混合物を20℃で1時間撹拌した。反応混合物を飽和塩化アンモニウム水溶液(150mL)によってクエンチし、酢酸エチル(100mL)で2回抽出した。0℃でpH=3になるまで、水層を塩酸水溶液(1M)によって調整し、次いで、クロロホルムおよびイソプロパノール(v/v=5:1、150mL、3回)で抽出した。合わせた層を無水硫酸ナトリウムで乾燥させ、濾過し、濾液を濃縮して、粗生成物を得た。化合物2-(4-フルオロ-3-テトラヒドロピラン-2-イルオキシ-イソキサゾール-5-イル)酢酸(8.07g、32.91mmol)を黄色の固体として得た。 To a solution of 4-fluoro-5-methyl-3-tetrahydropyran-2-yloxy-isoxazole (8.07 g, 40.13 mmol, 1 eq.) in tetrahydrofuran (115 mL), potassium bis(trimethylsilyl)amide (1 M, 60.20 mL, 1.5 eq.) was added dropwise under nitrogen at −78° C. and the reaction mixture was stirred at −78° C. for 0.5 h. Carbon dioxide (15 psi) was then bubbled through the reaction mixture at −78° C. for 1 h and the mixture was stirred at 20° C. for 1 h. The reaction mixture was quenched with saturated aqueous ammonium chloride solution (150 mL) and extracted twice with ethyl acetate (100 mL). The aqueous layer was adjusted to pH=3 at 0° C. with aqueous hydrochloric acid (1 M) and then extracted with chloroform and isopropanol (v/v=5:1, 150 mL, 3 times). The combined layers were dried over anhydrous sodium sulfate, filtered, and the filtrate was concentrated to give the crude product. Compound 2-(4-fluoro-3-tetrahydropyran-2-yloxy-isoxazol-5-yl)acetic acid (8.07 g, 32.91 mmol) was obtained as a yellow solid.
工程6

メタノール(70mL)および酢酸エチル(70mL)の混合物中の2-(4-フルオロ-3-テトラヒドロピラン-2-イルオキシ-イソキサゾール-5-イル)酢酸(7.87g、32.10mmol、1当量)の溶液に、トリメチルシリルジアゾメタンエーテル溶液(2M、24.07mL、1.5当量)を-10℃で、滴下で添加し、混合物を-10℃で1時間撹拌した。反応混合物を酢酸でpH=7まで調整し、酢酸エチル(150mL)を添加した。有機層を飽和重炭酸ナトリウム溶液(50mL)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、濾液を濃縮して、粗生成物を得た。粗生成物をシリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=1:0~5:1)によって精製した。化合物メチル2-(4-フルオロ-3-テトラヒドロピラン-2-イルオキシ-イソキサゾール-5-イル)酢酸塩(4.51g、17.40mmol、収率54%)を無色の油として得た。 To a solution of 2-(4-fluoro-3-tetrahydropyran-2-yloxy-isoxazol-5-yl)acetic acid (7.87 g, 32.10 mmol, 1 eq.) in a mixture of methanol (70 mL) and ethyl acetate (70 mL), trimethylsilyl diazomethane ether solution (2 M, 24.07 mL, 1.5 eq.) was added dropwise at -10 °C, and the mixture was stirred at -10 °C for 1 h. The reaction mixture was adjusted to pH = 7 with acetic acid, and ethyl acetate (150 mL) was added. The organic layer was washed with saturated sodium bicarbonate solution (50 mL), dried over anhydrous sodium sulfate, filtered, and the filtrate was concentrated to give the crude product. The crude product was purified by silica gel chromatography (petroleum ether: ethyl acetate = 1:0 to 5:1). The compound methyl 2-(4-fluoro-3-tetrahydropyran-2-yloxy-isoxazol-5-yl)acetate (4.51 g, 17.40 mmol, yield 54%) was obtained as a colorless oil.
工程7

ジメチルホルムアミド(66mL)中のメチル2-(4-フルオロ-3-テトラヒドロピラン-2-イルオキシ-イソキサゾール-5-イル)酢酸塩(4.42g、17.05mmol、1当量)の溶液に、tert-ブトキシドカリウム(2.87g、25.58mmol、1.5当量)を0℃で添加し、次いで、2-ヨードプロパン(3.48g、20.46mmol、2.05mL、1.2当量)を添加した。混合物を20℃で12時間撹拌した。反応混合物を水(60mL)で希釈し、酢酸エチル(80mL)で3回抽出した。合わせた層を飽和塩化ナトリウム水溶液(100mL)で2回洗浄し、無水硫酸ナトリウムで乾燥させ、濃縮して、粗生成物を得た。粗生成物を、シリカゲルクロマトグラフィー(石油エーテル)によって精製した。化合物メチル2-(4-フルオロ-3-テトラヒドロピラン-2-イルオキシ-イソキサゾール-5-イル)-3-メチル-ブタノエート(5.25g)を黄色の油として得た。 To a solution of methyl 2-(4-fluoro-3-tetrahydropyran-2-yloxy-isoxazol-5-yl)acetate (4.42 g, 17.05 mmol, 1 equiv.) in dimethylformamide (66 mL) was added potassium tert-butoxide (2.87 g, 25.58 mmol, 1.5 equiv.) at 0°C, followed by 2-iodopropane (3.48 g, 20.46 mmol, 2.05 mL, 1.2 equiv.). The mixture was stirred at 20°C for 12 h. The reaction mixture was diluted with water (60 mL) and extracted three times with ethyl acetate (80 mL). The combined layers were washed twice with saturated aqueous sodium chloride solution (100 mL), dried over anhydrous sodium sulfate, and concentrated to give the crude product. The crude product was purified by silica gel chromatography (petroleum ether). The compound methyl 2-(4-fluoro-3-tetrahydropyran-2-yloxy-isoxazol-5-yl)-3-methyl-butanoate (5.25 g) was obtained as a yellow oil.
工程8
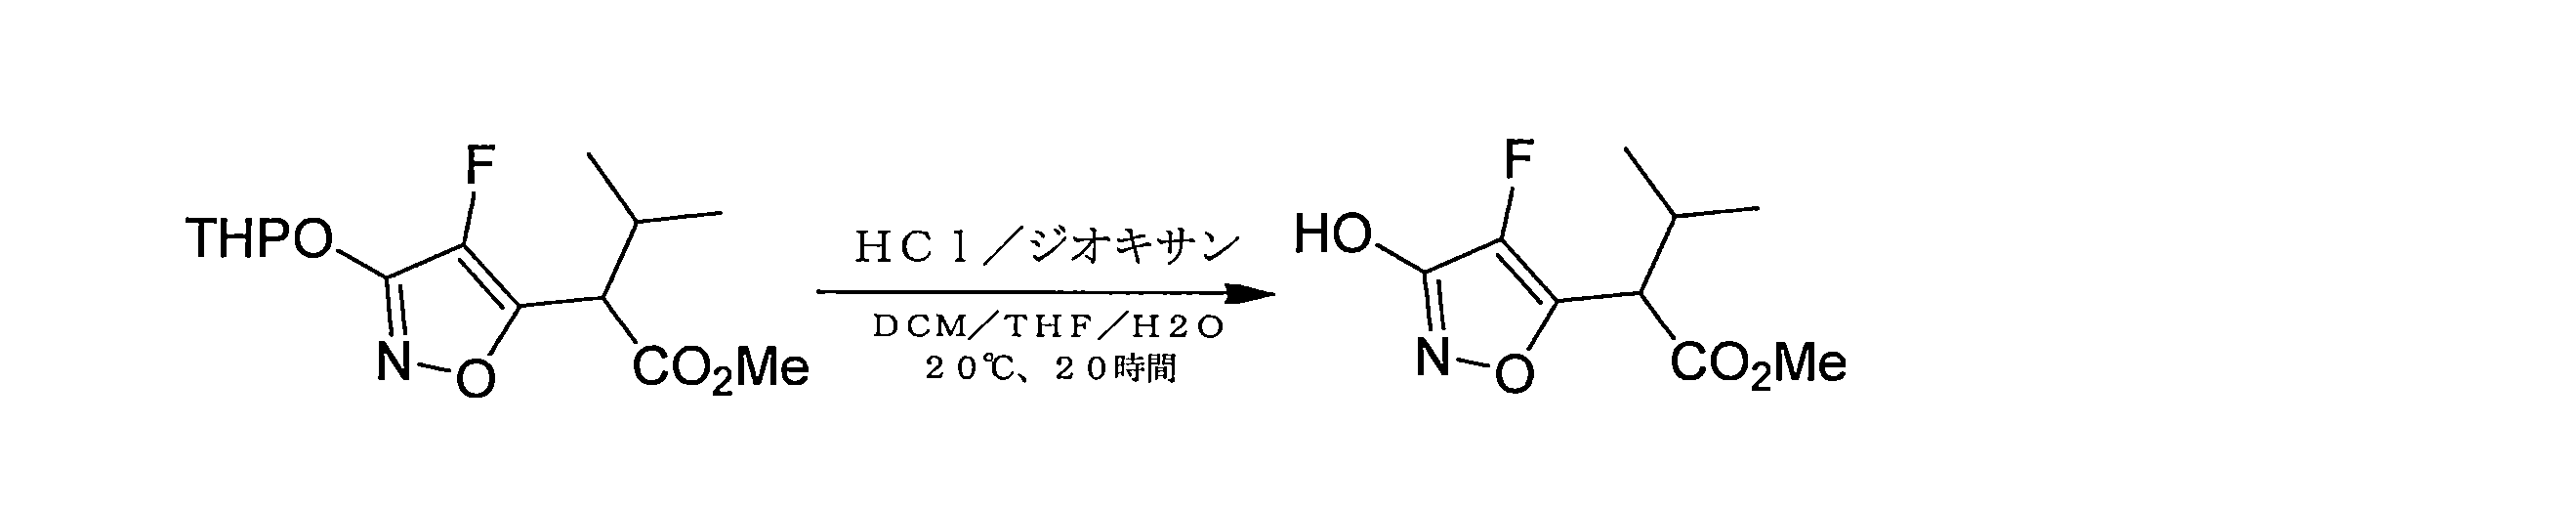
ジクロロメタン(20mL)およびテトラヒドロフラン(5mL)および水(5mL)の混合物中のメチル2-(4-フルオロ-3-テトラヒドロピラン-2-イルオキシ-イソキサゾール-5-イル)-3-メチル-ブタノエート(5.48g、18.19mmol、1当量)の溶液に、塩酸/ジオキサン(4M、30.00mL、6.60当量)を添加した。混合物を20℃で20時間撹拌した。反応混合物を飽和炭酸ナトリウム溶液(300mL)に注ぎ入れ、次いで、メチルtert-ブチルエーテル(100mL)で3回抽出した。水層のpHを、pH=2になるまで塩酸溶液(3M)を添加することによって調整した。水層をクロロホルム(150mL)で3回抽出し、無水硫酸ナトリウムで乾燥させ、濃縮して、粗生成物を得た。残渣を、半分取逆相(カラム:Kromasil 250*50mm*10um;移動相:[水(0.225%FA)-ACN];B%:21ACN%~51ACN%、29分、79%分)によって精製した。化合物メチル2-(4-フルオロ-3-ヒドロキシ-イソキサゾール-5-イル)-3-メチル-ブタノエート(1.27g、5.86mmol、収率32%)を白色の固体として得た。 To a solution of methyl 2-(4-fluoro-3-tetrahydropyran-2-yloxy-isoxazol-5-yl)-3-methyl-butanoate (5.48 g, 18.19 mmol, 1 eq.) in a mixture of dichloromethane (20 mL) and tetrahydrofuran (5 mL) and water (5 mL), hydrochloric acid/dioxane (4 M, 30.00 mL, 6.60 eq.) was added. The mixture was stirred at 20 °C for 20 h. The reaction mixture was poured into saturated sodium carbonate solution (300 mL) and then extracted three times with methyl tert-butyl ether (100 mL). The pH of the aqueous layer was adjusted by adding hydrochloric acid solution (3 M) until pH = 2. The aqueous layer was extracted three times with chloroform (150 mL), dried over anhydrous sodium sulfate and concentrated to give the crude product. The residue was purified by semi-preparative reverse phase (column: Kromasil 250*50mm*10um; mobile phase: [water (0.225% FA)-ACN]; B%: 21ACN% to 51ACN%, 29 min, 79% min). The compound methyl 2-(4-fluoro-3-hydroxy-isoxazol-5-yl)-3-methyl-butanoate (1.27g, 5.86mmol, 32% yield) was obtained as a white solid.
工程9

メチル2-(4-フルオロ-3-ヒドロキシ-イソキサゾール-5-イル)-3-メチル-ブタノエート(0.5g、2.30mmol、1当量)、2-ブロモ-1,1-ジエトキシ-エタン(907mg、4.60mmol、0.69mL、2当量)および炭酸カリウム(636mg、4.60mmol、2当量)を、ジメチルスルホキシド(8mL)中のマイクロ波チューブに取り込んだ。密封チューブを、マイクロ波照射下で、80℃で2時間加熱した。LCMSが出発材料が依然として残っていることを示した後、密封チューブを、マイクロ波照射下で、80℃でさらに2時間加熱した。混合物を濾過し、真空中で濃縮した。残渣を、半分取逆相(カラム:Phenomenex Synergi C18 150*25*10um;移動相:[水(0.225%FA)-ACN];B%:40%~70%、9分)によって精製した。化合物メチル2-[3-(2,2-ジエトキシエトキシ)-4-フルオロ-イソキサゾール-5-イル]-3-メチル-ブタノエート(266mg、0.80mmol、収率35%)を黄色の油として得、46mgのメチル2-(4-フルオロ-3-ヒドロキシ-イソキサゾール-5-イル)-3-メチル-ブタノエートを回収した。 Methyl 2-(4-fluoro-3-hydroxy-isoxazol-5-yl)-3-methyl-butanoate (0.5 g, 2.30 mmol, 1 eq.), 2-bromo-1,1-diethoxy-ethane (907 mg, 4.60 mmol, 0.69 mL, 2 eq.) and potassium carbonate (636 mg, 4.60 mmol, 2 eq.) were taken in a microwave tube in dimethylsulfoxide (8 mL). The sealed tube was heated under microwave irradiation at 80 °C for 2 h. After LCMS showed starting material still remained, the sealed tube was heated under microwave irradiation at 80 °C for an additional 2 h. The mixture was filtered and concentrated in vacuo. The residue was purified by semi-preparative reverse phase (column: Phenomenex Synergi C18 150*25*10um; mobile phase: [water (0.225% FA)-ACN]; B%: 40%-70%, 9 min). Compound methyl 2-[3-(2,2-diethoxyethoxy)-4-fluoro-isoxazol-5-yl]-3-methyl-butanoate (266 mg, 0.80 mmol, 35% yield) was obtained as a yellow oil, and 46 mg of methyl 2-(4-fluoro-3-hydroxy-isoxazol-5-yl)-3-methyl-butanoate was recovered.
例示的な化合物76について上に記載されるものと同様の手順を使用して、メチル2-[3-(2,2-ジエトキシエトキシ)-4-フルオロ-イソキサゾール-5-イル]-3-メチル-ブタノエートを最終化合物に変換した。 Using a procedure similar to that described above for exemplary compound 76, methyl 2-[3-(2,2-diethoxyethoxy)-4-fluoro-isoxazol-5-yl]-3-methyl-butanoate was converted to the final compound.
例示的な化合物198、208、209、269、および270を、類似の手順を使用して調製した。 Exemplary compounds 198, 208, 209, 269, and 270 were prepared using similar procedures.
例示的な化合物200の例示的な合成

工程1
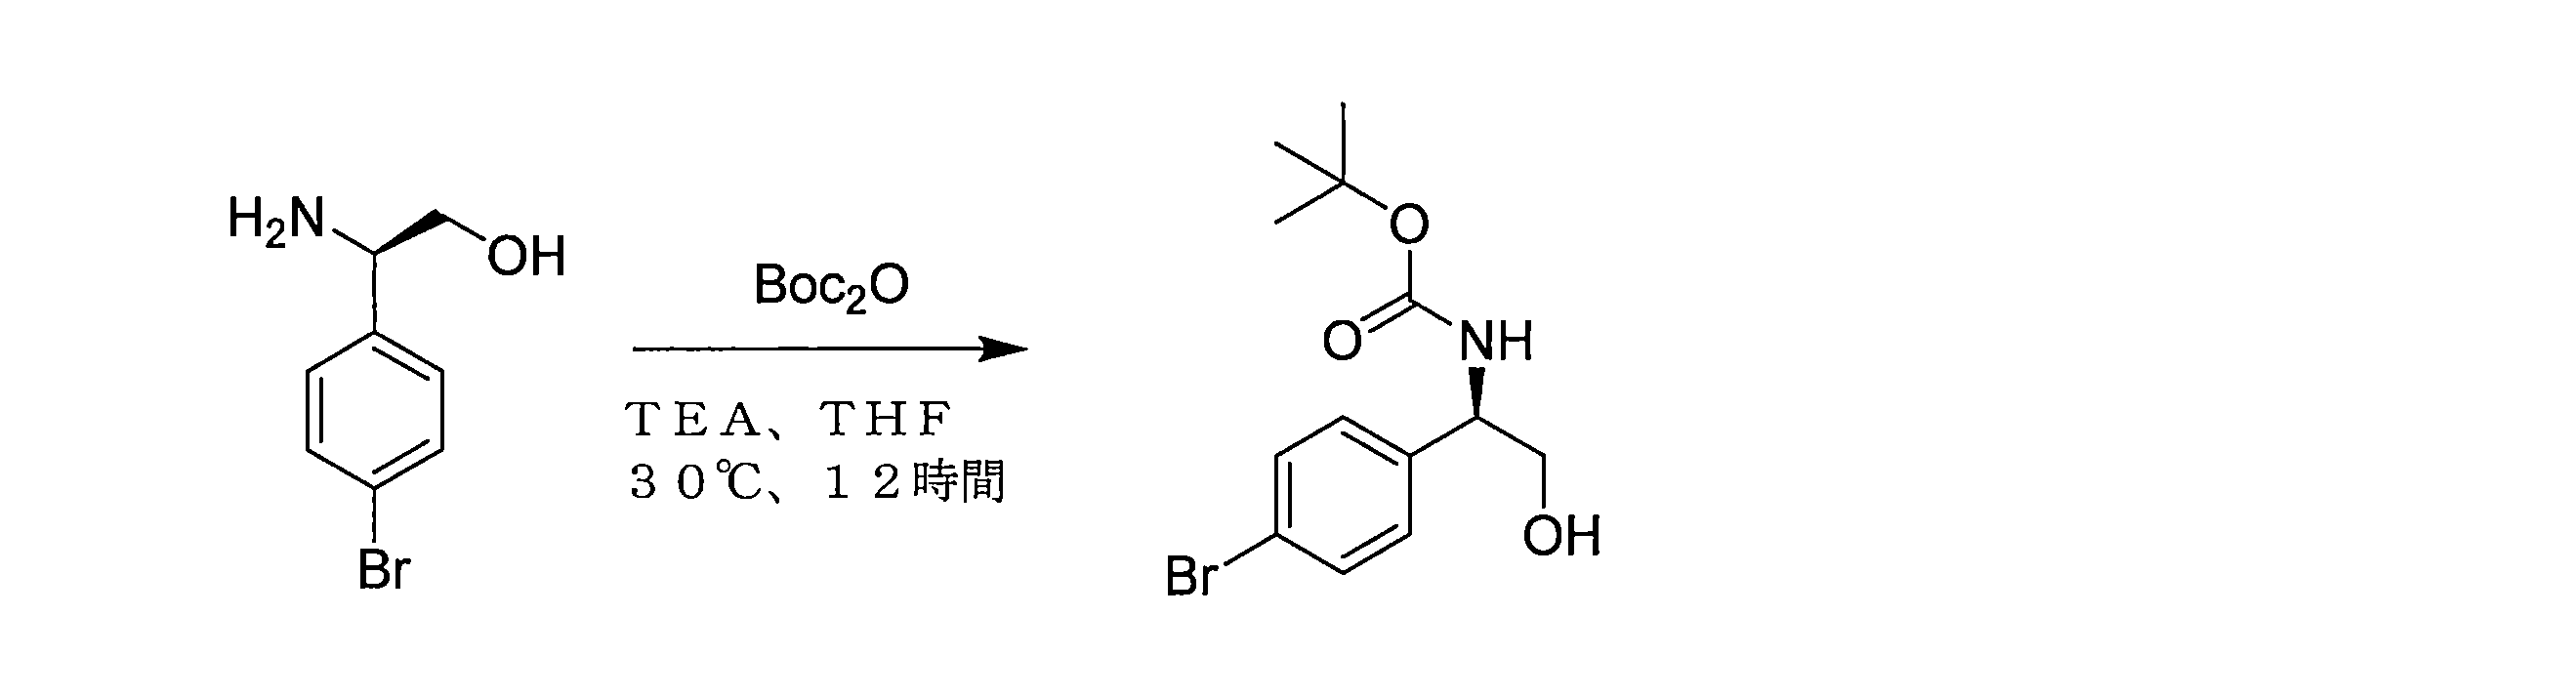
(2R)-2-アミノ-2-(4-ブロモフェニル)エタノール(1.74g、6.89mmol、1当量、塩酸塩)のテトラヒドロフラン(25mL)の溶液に、トリエチルアミン(2.79g、27.56mmol、3.84mL、4当量)およびtert-ブトキシカルボニルtert-ブチル炭酸塩(1.80g、8.27mmol、1.90mL、1.2当量)を加えた。混合物を30℃で12時間撹拌した。反応混合物を減圧下で濃縮して、テトラヒドロフランを除去した。残渣を酢酸エチル(50mL×2)で抽出した。合わせた有機層をブライン水溶液(40mL×2)で洗浄し、無水硫酸ナトリウム上で乾燥させ、濾過し、減圧下で濃縮した。粗生成物を、石油エーテル/酢酸エチル(22mL、v:v=10:1)で粉砕した。化合物tert-ブチルN-[(1R)-1-(4-ブロモフェニル)-2-ヒドロキシ-エチル]カルバメート(2g、6.33mmol、収率91%)を白色の固体として得た。 To a solution of (2R)-2-amino-2-(4-bromophenyl)ethanol (1.74 g, 6.89 mmol, 1 eq., hydrochloride salt) in tetrahydrofuran (25 mL) was added triethylamine (2.79 g, 27.56 mmol, 3.84 mL, 4 eq.) and tert-butoxycarbonyl tert-butyl carbonate (1.80 g, 8.27 mmol, 1.90 mL, 1.2 eq.). The mixture was stirred at 30°C for 12 h. The reaction mixture was concentrated under reduced pressure to remove tetrahydrofuran. The residue was extracted with ethyl acetate (50 mL x 2). The combined organic layers were washed with aqueous brine solution (40 mL x 2), dried over anhydrous sodium sulfate, filtered and concentrated under reduced pressure. The crude product was triturated with petroleum ether/ethyl acetate (22 mL, v:v=10:1). The compound tert-butyl N-[(1R)-1-(4-bromophenyl)-2-hydroxy-ethyl]carbamate (2 g, 6.33 mmol, 91% yield) was obtained as a white solid.
工程2
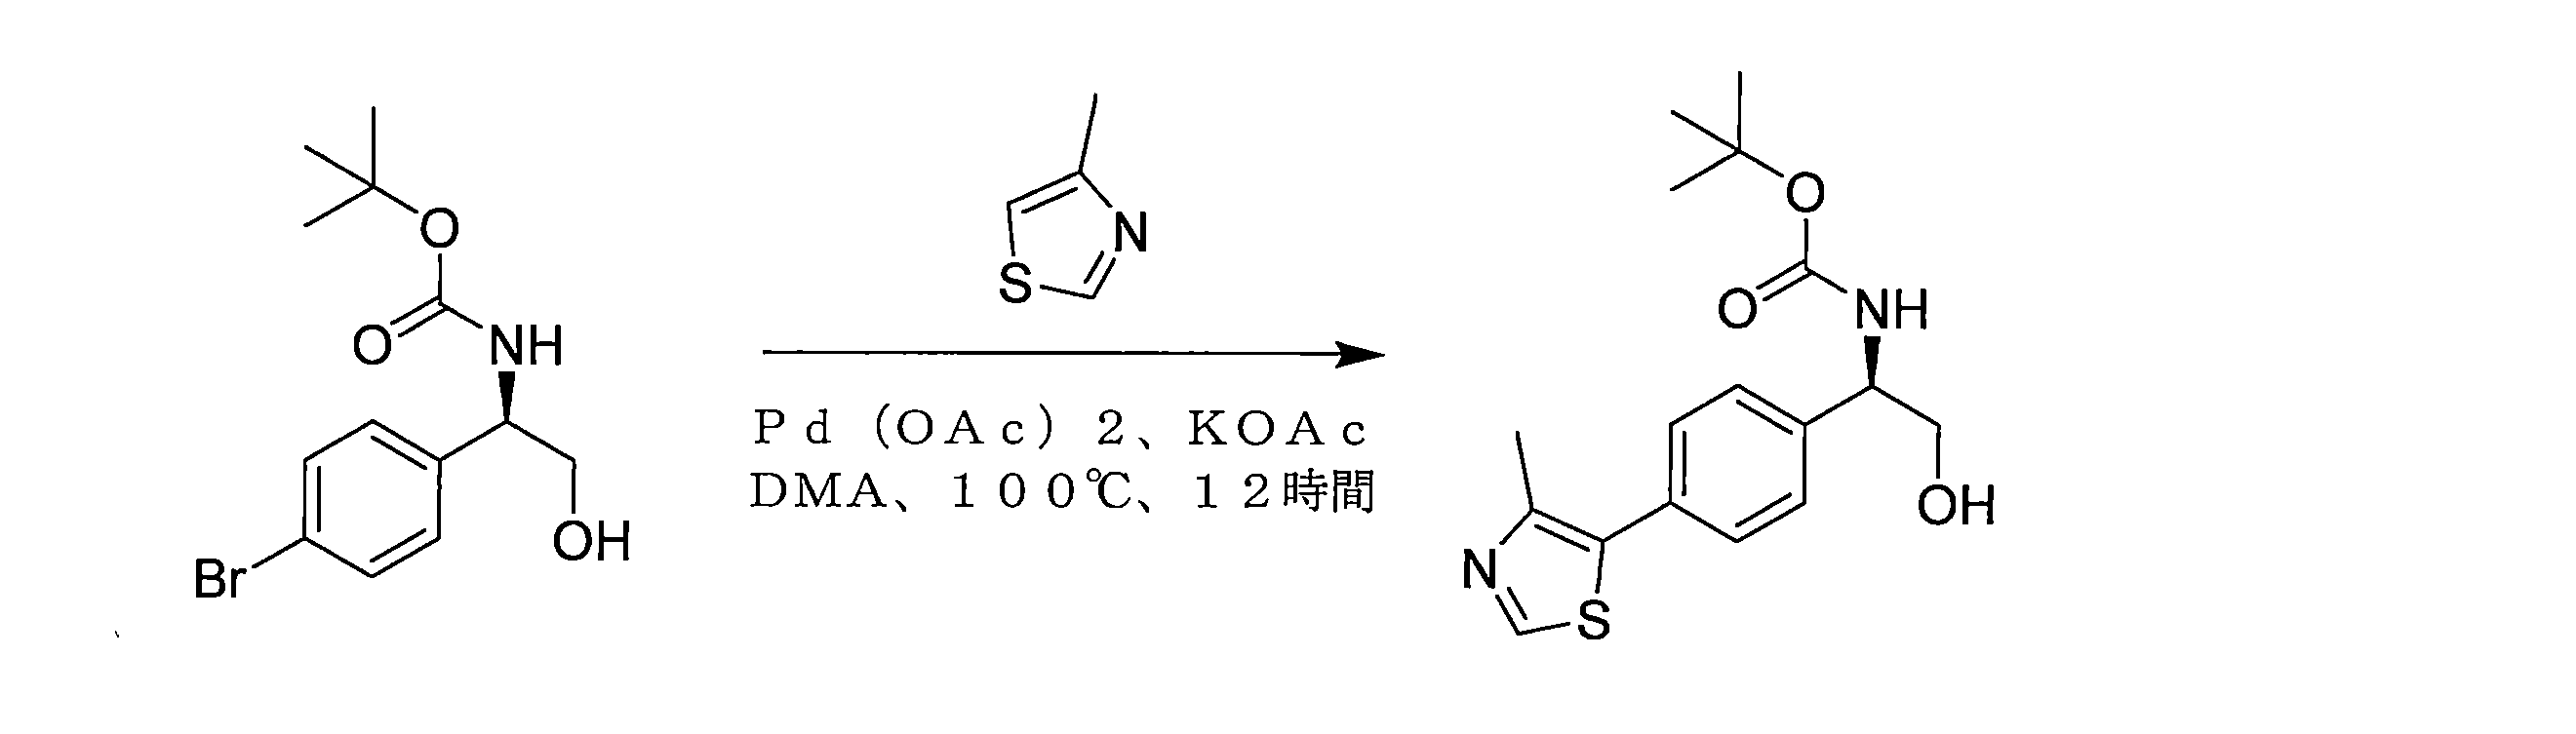
ジメチルアセトアミド(40mL)中のtert-ブチルN-[(1R)-1-(4-ブロモフェニル)-2-ヒドロキシ-エチル]カルバメート(2.0g、6.33mmol、1当量)、4-メチルチアゾール(1.25g、12.65mmol、1.15mL、2当量)、酢酸カリウム(1.24g、12.65mmol、2当量)、および酢酸パラジウム(142mg、0.63mmol、0.1当量)の混合物を、脱気し、窒素で3回パージし、次いで、混合物を窒素雰囲気下で、100℃で12時間撹拌した。反応混合物を減圧下で濃縮して、ジメチルアセトアミドを除去した。残渣を酢酸エチル(40mL×2)で抽出し、飽和ブライン(40mL×2)で洗浄した。合わせた有機層を無水硫酸ナトリウム上で乾燥させ、濾過し、減圧下で濃縮した。残渣を、シリカゲルクロマトグラフィー(石油エーテル/酢酸エチル=5/1~1/2)によって精製した。化合物tert-ブチルN-[(1R)-2-ヒドロキシ-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]カルバメート(1.45g、3.98mmol、収率63%)を黄色の油として得た。 A mixture of tert-butyl N-[(1R)-1-(4-bromophenyl)-2-hydroxy-ethyl]carbamate (2.0 g, 6.33 mmol, 1 eq.), 4-methylthiazole (1.25 g, 12.65 mmol, 1.15 mL, 2 eq.), potassium acetate (1.24 g, 12.65 mmol, 2 eq.), and palladium acetate (142 mg, 0.63 mmol, 0.1 eq.) in dimethylacetamide (40 mL) was degassed and purged with nitrogen three times, and then the mixture was stirred at 100°C under nitrogen atmosphere for 12 hours. The reaction mixture was concentrated under reduced pressure to remove dimethylacetamide. The residue was extracted with ethyl acetate (40 mL x 2) and washed with saturated brine (40 mL x 2). The combined organic layers were dried over anhydrous sodium sulfate, filtered, and concentrated under reduced pressure. The residue was purified by silica gel chromatography (petroleum ether/ethyl acetate = 5/1 to 1/2). The compound tert-butyl N-[(1R)-2-hydroxy-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]carbamate (1.45 g, 3.98 mmol, 63% yield) was obtained as a yellow oil.
上記の例示的な化合物180について記載されるものと類似の手順を使用して、tert-ブチルN-[(1R)-2-ヒドロキシ-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]カルバメートを最終化合物に変換した。 Using a procedure similar to that described for exemplary compound 180 above, tert-butyl N-[(1R)-2-hydroxy-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]carbamate was converted to the final compound.
例示的な化合物199、158、および100を、類似の手順を使用して調製した。 Exemplary compounds 199, 158, and 100 were prepared using similar procedures.
例示的な化合物202の例示的な合成

工程1

ジメチルホルムアミド(50mL)中の1-(4-ブロモ-2-ヒドロキシ-フェニル)エタノン(10g、46.50mmol、1.0当量)の溶液に、炭酸カリウム(9.64g、69.75mmol、1.5当量)を添加した。次いで、ヨードメタン(13.20g、93.00mmol、2.0当量)を0℃で混合物中に添加した。次いで、混合物を20℃で12時間撹拌した。混合物を水(200mL)で希釈し、酢酸エチル(100mL×3)で抽出し、ブライン(50mL×2)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、次いで、濃縮した。1-(4-ブロモ-2-メトキシ-フェニル)エタノン(10.5g、45.84mmol、収率98%)を黄色の固体として得た。 To a solution of 1-(4-bromo-2-hydroxy-phenyl)ethanone (10 g, 46.50 mmol, 1.0 equiv.) in dimethylformamide (50 mL) was added potassium carbonate (9.64 g, 69.75 mmol, 1.5 equiv.). Then, iodomethane (13.20 g, 93.00 mmol, 2.0 equiv.) was added into the mixture at 0°C. The mixture was then stirred at 20°C for 12 hours. The mixture was diluted with water (200 mL), extracted with ethyl acetate (100 mL x 3), washed with brine (50 mL x 2), dried over anhydrous sodium sulfate, filtered, and then concentrated. 1-(4-bromo-2-methoxy-phenyl)ethanone (10.5 g, 45.84 mmol, 98% yield) was obtained as a yellow solid.
工程2
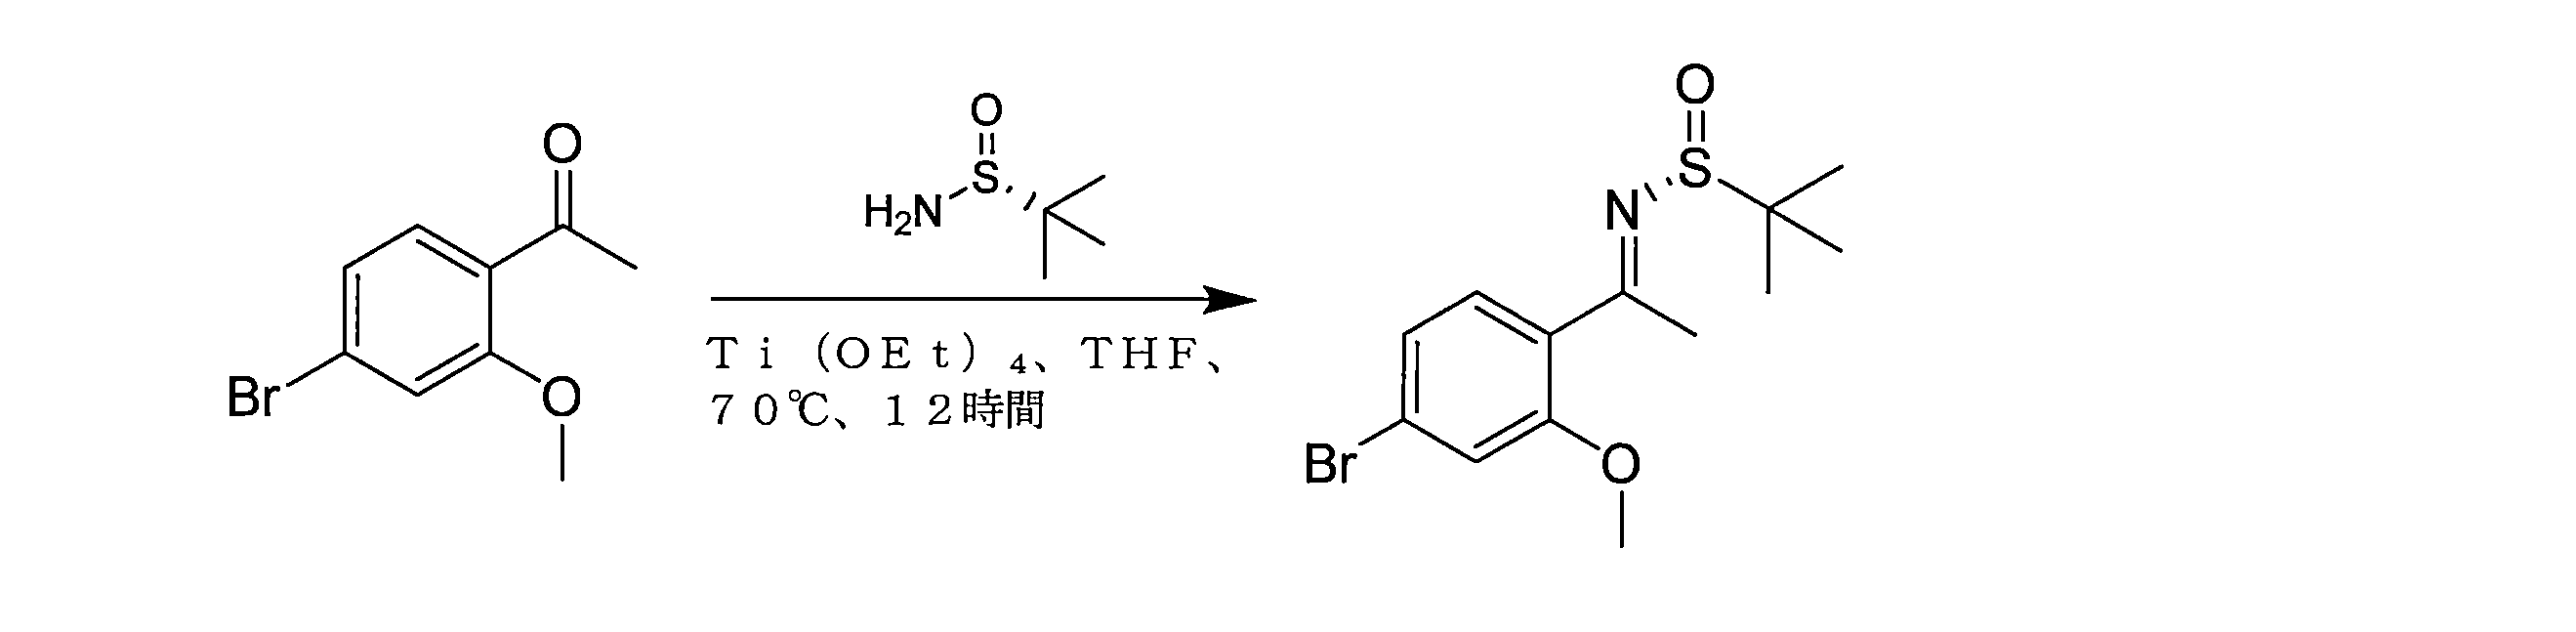
テトラヒドロフラン(50mL)中の1-(4-ブロモ-2-メトキシ-フェニル)エタノン(10g、43.65mmol、1.18当量)の溶液に、チタンテトラエトキシド(16.94g、74.26mmol、2.0当量)を添加した。次いで、2-メチルプロパン-2-スルフィンアミド(4.5g、37.13mmol、1.0当量)を混合物に添加し、混合物をN2で3回パージした。次いで、混合物を70℃で12時間撹拌した。混合物を水(100mL)でクエンチし、水(200mL)で希釈し、濾過し、次いで、酢酸エチル(200mL×3)で抽出し、ブライン(200mL)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、次いで、濃縮した。残渣を、シリカゲルカラムクロマトグラフィー(石油エーテル:酢酸エチル=20:1~3:1)によって精製した。((R,E)-N-(1-(4-ブロモ-2-メトキシフェニル)エチリデン)-2-メチルプロパン-2-スルフィンアミド(9g、27.09mmol、収率73%)を黄色の油として得た。 To a solution of 1-(4-bromo-2-methoxy-phenyl)ethanone (10 g, 43.65 mmol, 1.18 eq.) in tetrahydrofuran (50 mL), titanium tetraethoxide (16.94 g, 74.26 mmol, 2.0 eq.) was added. Then, 2-methylpropane-2-sulfinamide (4.5 g, 37.13 mmol, 1.0 eq.) was added to the mixture, and the mixture was purged with N2 three times. The mixture was then stirred at 70°C for 12 hours. The mixture was quenched with water (100 mL), diluted with water (200 mL), filtered, then extracted with ethyl acetate (200 mL x 3), washed with brine (200 mL), dried over anhydrous sodium sulfate, filtered, and then concentrated. The residue was purified by silica gel column chromatography (petroleum ether:ethyl acetate = 20:1 to 3:1). ((R,E)-N-(1-(4-bromo-2-methoxyphenyl)ethylidene)-2-methylpropane-2-sulfinamide (9 g, 27.09 mmol, 73% yield) was obtained as a yellow oil.
工程3

テトラヒドロフラン(90mL)中の((R,E)-N-(1-(4-ブロモ-2-メトキシフェニル)エチリデン)-2-メチルプロパン-2-スルフィンアミド(9g、27.09mmol、1.0当量)の溶液に、L-セレクトリド(1M、81.26mL、3.0当量)を0℃で添加した。次いで、混合物を20℃で2時間撹拌した。混合物を水(100mL)でクエンチし、水(20mL)で希釈し、酢酸エチル(300mL×3)で抽出し、ブライン(200mL)で洗浄し、無水硫酸ナトリウム上で乾燥させ、濾過し、次いで濃縮した。残渣を、シリカゲルカラムクロマトグラフィー(石油エーテル:酢酸エチル=20:1~1:1)によって精製した。N-[(1S)-1-(4-ブロモ-2-メトキシ-フェニル)エチル]-2-メチル-プロパン-2-スルフィンアミド(5.5g、16.45mmol、収率60%)を黄色の油として得た。 To a solution of ((R,E)-N-(1-(4-bromo-2-methoxyphenyl)ethylidene)-2-methylpropane-2-sulfinamide (9 g, 27.09 mmol, 1.0 equiv.) in tetrahydrofuran (90 mL) was added L-selectride (1 M, 81.26 mL, 3.0 equiv.) at 0° C. The mixture was then stirred at 20° C. for 2 h. The mixture was quenched with water (100 mL), diluted with water (20 mL), and diluted with ethyl acetate (300 mL). The mixture was extracted with 100 mL of hexane (3×100 mL), washed with brine (200 mL), dried over anhydrous sodium sulfate, filtered, and then concentrated. The residue was purified by silica gel column chromatography (petroleum ether:ethyl acetate=20:1 to 1:1). N-[(1S)-1-(4-bromo-2-methoxy-phenyl)ethyl]-2-methyl-propane-2-sulfinamide (5.5 g, 16.45 mmol, 60% yield) was obtained as a yellow oil.
工程4

ジオキサン(12mL)中のN-[(1S)-1-(4-ブロモ-2-メトキシ-フェニル)エチル]-2-メチル-プロパン-2-スルフィンアミド(4.7g、14.06mmol、1.0当量)および4,4,5,5-テトラメチル-2-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)-1,3,2-ジオキサボロラン(5.36g、21.09mmol、1.5当量)の溶液に、[1,1’-ビス(ジフェニルホスフィノ)フェロセン]ジクロロパラジウム(ii)(1.03g、1.41mmol、0.1当量)および酢酸カリウム(2.76g、28.12mmol、2.0当量)を添加した。次いで、混合物をN2で3回パージした。次いで、混合物を90℃で5時間撹拌した。混合物を水(20mL)で希釈し、濾過し、次いで、酢酸エチル(50mL×3)で抽出し、ブライン(30mL)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、次いで、濃縮した。残渣を、シリカゲルカラムクロマトグラフィー(石油エーテル:酢酸エチル=10:1~1:1)によって精製した。N-[(1S)-1-[2-メトキシ-4-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)フェニル]エチル]-2-メチル-プロパン-2-スルフィンアミド(4.5g、11.80mmol、収率84%)を黄色の油として得た。 To a solution of N-[(1S)-1-(4-bromo-2-methoxy-phenyl)ethyl]-2-methyl-propane-2-sulfinamide (4.7 g, 14.06 mmol, 1.0 eq.) and 4,4,5,5-tetramethyl-2-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)-1,3,2-dioxaborolane (5.36 g, 21.09 mmol, 1.5 eq.) in dioxane (12 mL), [1,1'-bis(diphenylphosphino)ferrocene]dichloropalladium(ii) (1.03 g, 1.41 mmol, 0.1 eq.) and potassium acetate (2.76 g, 28.12 mmol, 2.0 eq.) were added. The mixture was then purged with N2 three times. The mixture was then stirred at 90 °C for 5 h. The mixture was diluted with water (20 mL), filtered, then extracted with ethyl acetate (50 mL x 3), washed with brine (30 mL), dried over anhydrous sodium sulfate, filtered, and then concentrated. The residue was purified by silica gel column chromatography (petroleum ether: ethyl acetate = 10: 1 to 1: 1). N-[(1S)-1-[2-methoxy-4-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)phenyl]ethyl]-2-methyl-propane-2-sulfinamide (4.5 g, 11.80 mmol, yield 84%) was obtained as a yellow oil.
以下のスキームに従って、N-[(1S)-1-[2-メトキシ-4-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)フェニル]エチル]-2-メチル-プロパン-2-スルフィンアミドを、(S)-1-(2-メトキシ-4-(4-メチルチアゾール-5-イル)フェニル)エタン-1-アミンに変換した。

上記の他の実施例(例えば、例示的な化合物64)について記載されるものと類似の手順を使用して、S)-1-(2-メトキシ-4-(4-メチルチアゾール-5-イル)フェニル)エタン-1-アミンを最終化合物に変換した。 S)-1-(2-Methoxy-4-(4-methylthiazol-5-yl)phenyl)ethan-1-amine was converted to the final compound using procedures similar to those described for other examples above (e.g., exemplary compound 64).
例示的な化合物201を、上に記載されるものと類似の手順を使用して調製した。 Exemplary compound 201 was prepared using procedures similar to those described above.
例示的な化合物203の例示的な合成
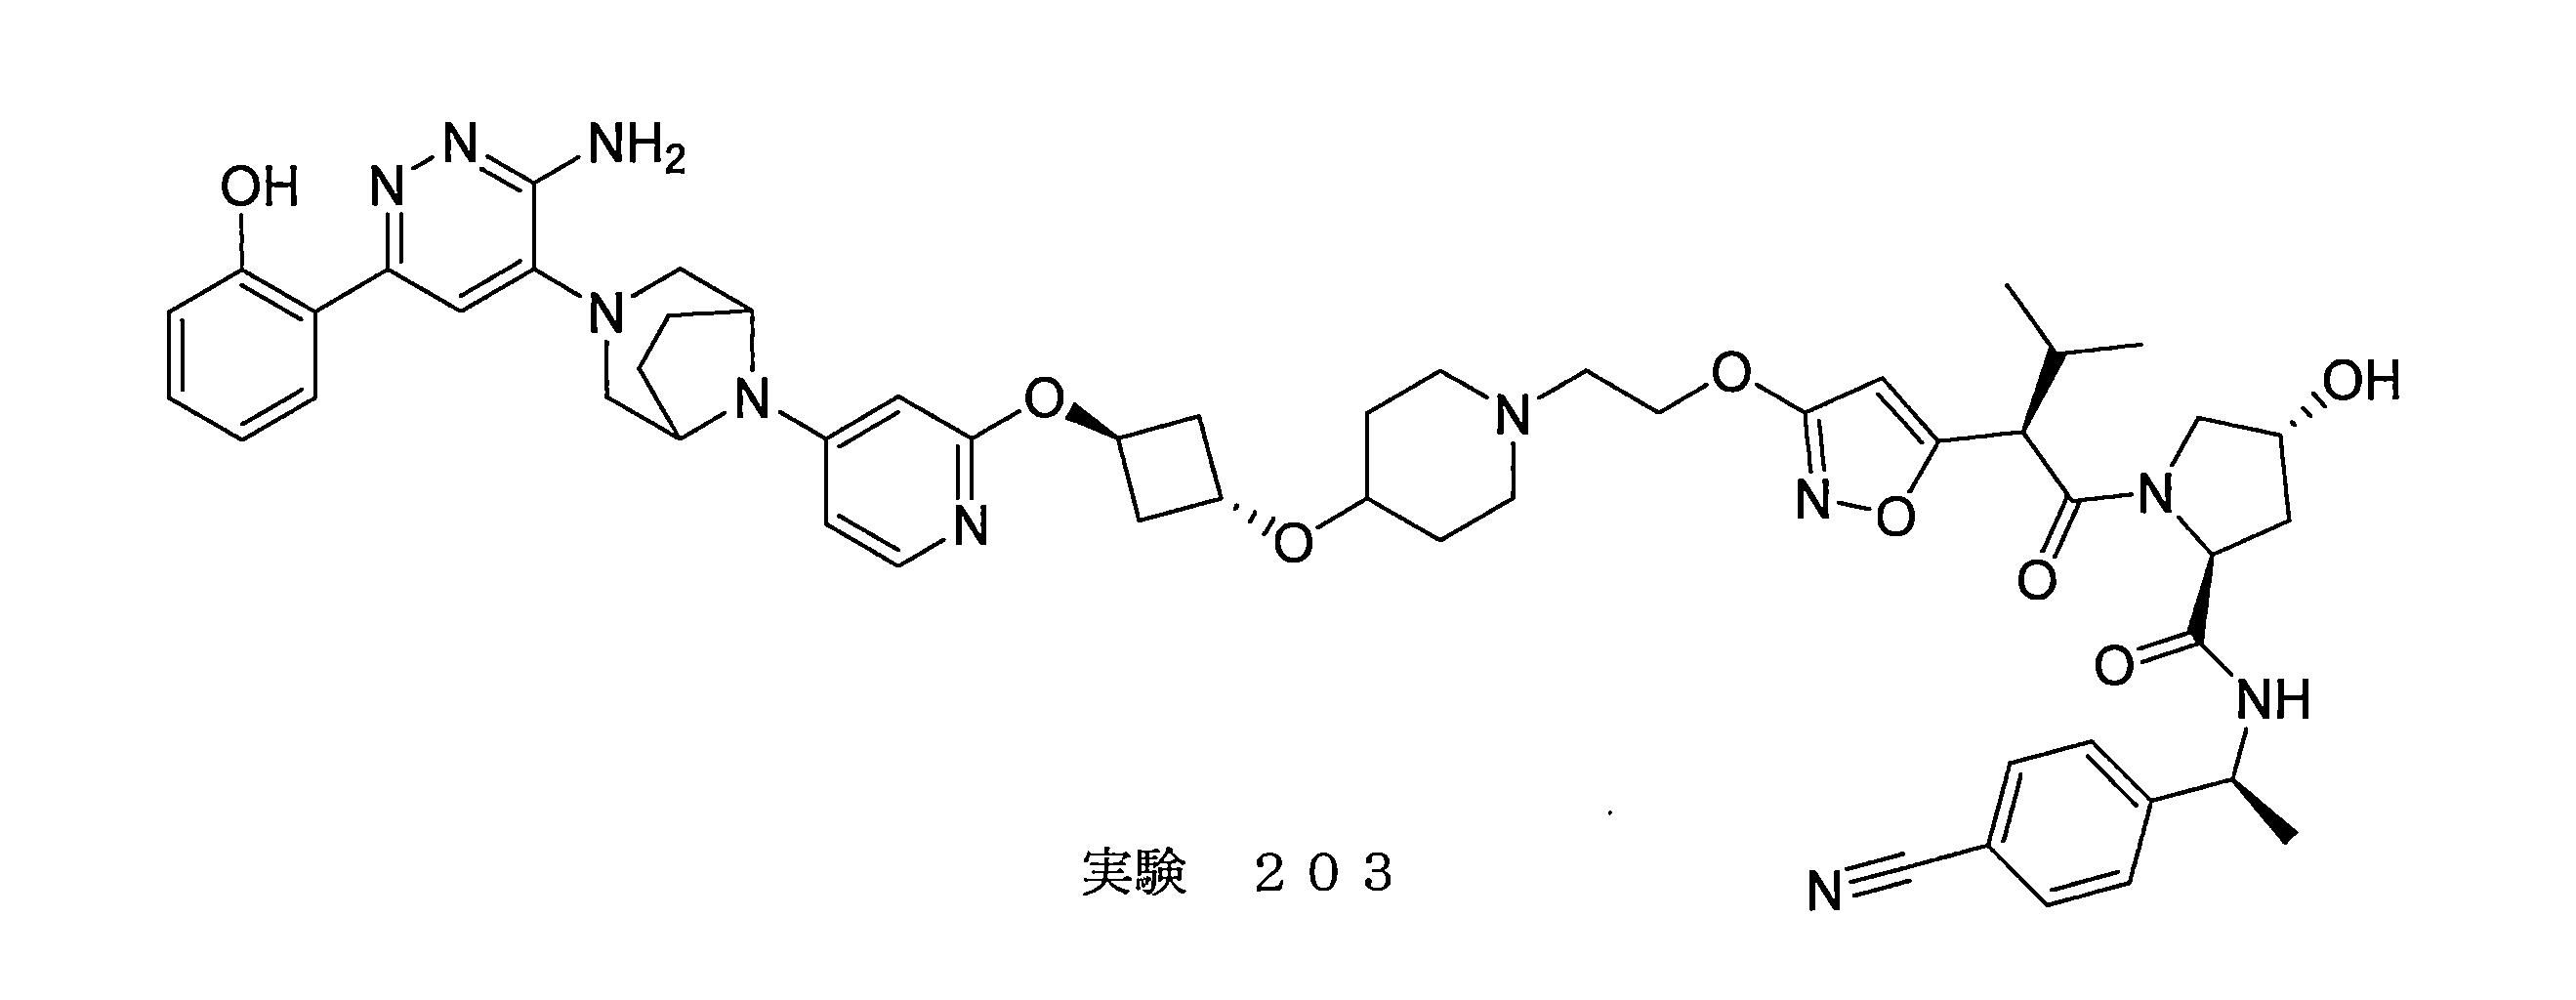
工程1

N,N-ジメチルホルムアミド(20mL)中のtert-ブチルN-[(1S)-1-(4-ブロモフェニル)エチル]カルバメート(2g、6.66mmol、1当量)の溶液に、パラジウムテトラキストリフェニルホスフィン(770mg、0.67mmol、0.1当量)およびシアン化亜鉛(782mg、6.66mmol、0.4mL、1当量)を添加した。反応溶液を80℃で14時間撹拌した。溶液を水(80mL)でクエンチし、酢酸エチル(50mL×3)で抽出した。合わせた有機層をブライン(20mL×3)で洗浄した。有機層を無水硫酸ナトリウムで乾燥させ、濾過し、減圧下で濃縮して、残渣を得た。残渣を、シリカゲルカラムクロマトグラフィー(石油エーテル/酢酸エチル=30/1~3:1)によって精製して、生成物を得た。tert-ブチルN-[(1S)-1-(4-シアノフェニル)エチル]カルバメート(1.6g、6.50mmol、収率97.50%)を無色のガムとして得た。 To a solution of tert-butyl N-[(1S)-1-(4-bromophenyl)ethyl]carbamate (2 g, 6.66 mmol, 1 eq.) in N,N-dimethylformamide (20 mL), palladium tetrakistriphenylphosphine (770 mg, 0.67 mmol, 0.1 eq.) and zinc cyanide (782 mg, 6.66 mmol, 0.4 mL, 1 eq.) were added. The reaction solution was stirred at 80° C. for 14 hours. The solution was quenched with water (80 mL) and extracted with ethyl acetate (50 mL×3). The combined organic layer was washed with brine (20 mL×3). The organic layer was dried over anhydrous sodium sulfate, filtered, and concentrated under reduced pressure to give a residue. The residue was purified by silica gel column chromatography (petroleum ether/ethyl acetate=30/1 to 3:1) to give the product. tert-Butyl N-[(1S)-1-(4-cyanophenyl)ethyl]carbamate (1.6 g, 6.50 mmol, 97.50% yield) was obtained as a colorless gum.
上に記載される手順を使用して、tert-ブチルN-[(1S)-1-(4-シアノフェニル)エチル]カルバメートを最終化合物に変換した。 Using the procedure described above, tert-butyl N-[(1S)-1-(4-cyanophenyl)ethyl]carbamate was converted to the final compound.
例示的な化合物204を、類似の手順を使用して調製した。 Exemplary compound 204 was prepared using a similar procedure.
例示的な化合物91の例示的な合成

工程1
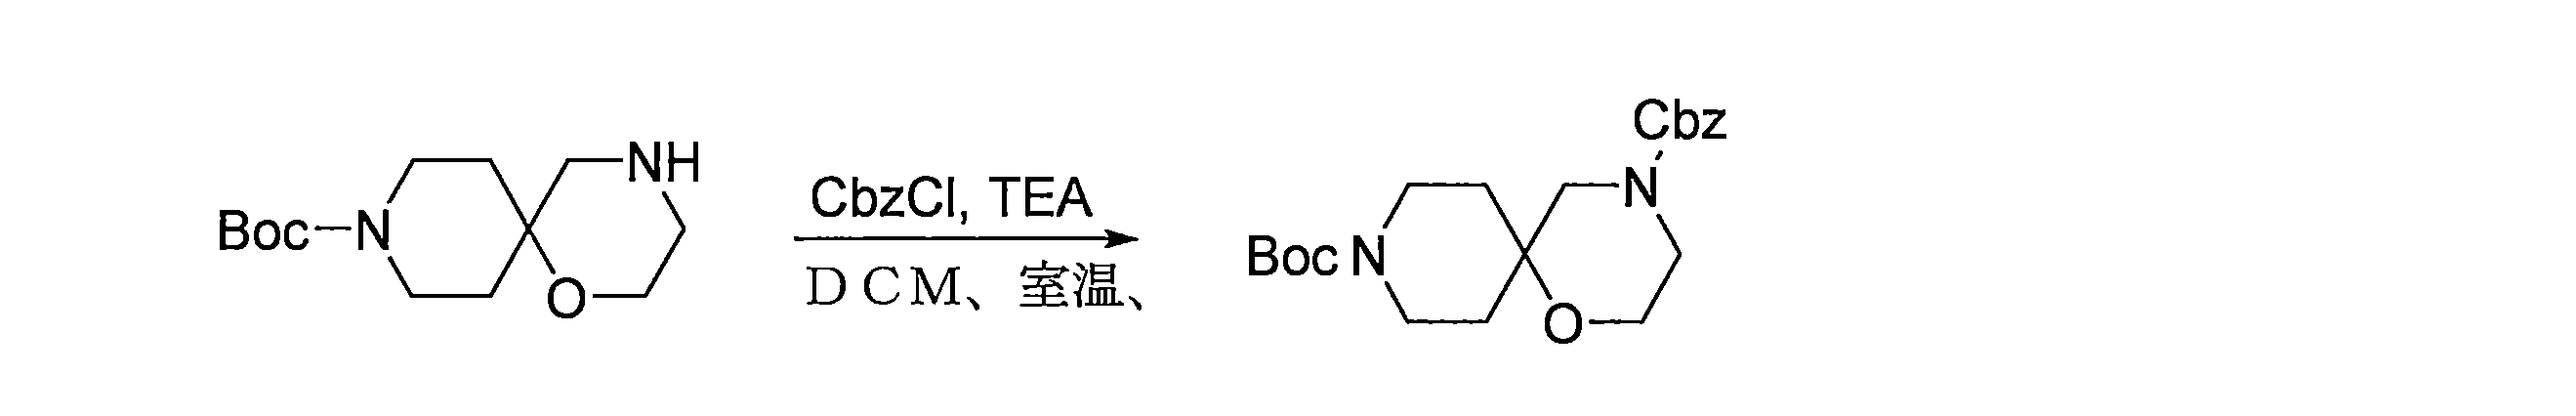
100mLの丸底フラスコに、DCM(8mL)中のtert-ブチル1-オキサ-4,9-ジアザスピロ[5.5]ウンデカン-9-カルボキシレート(3.5g、13.7mmol、1.0当量)およびTEA(4.1g、0.1mmol、3.0当量)を入れ、それにCbzCl(2.8g、0.1mmol、1.2当量)をゆっくり添加した。得られた混合物を、室温で12時間撹拌した。次いで、反応混合物を、20mLの水を添加することによってクエンチし、DCM(20mL×2)で抽出した。混合有機層を無水硫酸ナトリウム上で乾燥させ、減圧下で濃縮した。残渣を、酢酸エチル/石油エーテル(1:0)で溶出するシリカゲルカラム上に適用した。これにより、4.0g(75%)の4-ベンジル9-tert-ブチル1-オキサ-4,9-ジアザスピロ[5.5]ウンデカン-4,9-ジカルボキシレートを白色の油として得た。 A 100 mL round bottom flask was charged with tert-butyl 1-oxa-4,9-diazaspiro[5.5]undecane-9-carboxylate (3.5 g, 13.7 mmol, 1.0 equiv.) and TEA (4.1 g, 0.1 mmol, 3.0 equiv.) in DCM (8 mL), to which CbzCl (2.8 g, 0.1 mmol, 1.2 equiv.) was slowly added. The resulting mixture was stirred at room temperature for 12 h. The reaction mixture was then quenched by adding 20 mL of water and extracted with DCM (20 mL x 2). The combined organic layers were dried over anhydrous sodium sulfate and concentrated under reduced pressure. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (1:0). This yielded 4.0 g (75%) of 4-benzyl 9-tert-butyl 1-oxa-4,9-diazaspiro[5.5]undecane-4,9-dicarboxylate as a white oil.
工程2

100mLの丸底フラスコに、ジオキサン(15mL)中の4-ベンジル9-tert-ブチル1-オキサ-4,9-ジアザスピロ[5.5]ウンデカン-4,9-ジカルボキシレート(4.0g、10.2mmol、1.0当量)を入れ、次いで、1,4-ジオキサン溶液(4.0M、4mL)中の塩化水素を添加した。得られた混合物を室温で3時間撹拌し、次いで、減圧下で濃縮した。これにより、2.9g(87%)のベンジル1-オキサ-4,9-ジアザスピロ[5.5]ウンデカン-4-カルボキシレート塩酸塩を固体として得た。 A 100 mL round bottom flask was charged with 4-benzyl 9-tert-butyl 1-oxa-4,9-diazaspiro[5.5]undecane-4,9-dicarboxylate (4.0 g, 10.2 mmol, 1.0 equiv.) in dioxane (15 mL) and then hydrogen chloride in 1,4-dioxane solution (4.0 M, 4 mL) was added. The resulting mixture was stirred at room temperature for 3 hours and then concentrated under reduced pressure. This gave 2.9 g (87%) of benzyl 1-oxa-4,9-diazaspiro[5.5]undecane-4-carboxylate hydrochloride as a solid.
工程3

30mLの密封チューブに、DMSO(8mL)中のベンジル1-オキサ-4,9-ジアザスピロ[5.5]ウンデカン-4-カルボキシレート塩酸塩(2.9g、8.9mmol、1.0当量)、4-ブロモ-6-クロロピリダジン-3-アミン(2.1g、0.1mmol)、およびDIEA(3.9g、0.1mmol)を入れた。得られた混合物を、油浴中で、130℃で12時間撹拌した。次いで、反応混合物を、20mLの水/氷を添加することによってクエンチした。得られた混合物を酢酸エチル(20mL×2)で抽出した。混合有機層を無水硫酸ナトリウム上で乾燥させ、減圧下で濃縮した。残渣を、酢酸エチル/石油エーテル(1:0)で溶出するシリカゲルカラム上に適用した。これにより、3.1g(84%)のベンジル9-(3-アミノ-6-クロロピリダジン-4-イル)-1-オキサ-4,9-ジアザスピロ[5.5]ウンデカン-4-カルボキシレートを黄色の固体として得た。 A 30 mL sealed tube was charged with benzyl 1-oxa-4,9-diazaspiro[5.5]undecane-4-carboxylate hydrochloride (2.9 g, 8.9 mmol, 1.0 equiv.), 4-bromo-6-chloropyridazin-3-amine (2.1 g, 0.1 mmol), and DIEA (3.9 g, 0.1 mmol) in DMSO (8 mL). The resulting mixture was stirred in an oil bath at 130 °C for 12 h. The reaction mixture was then quenched by adding 20 mL of water/ice. The resulting mixture was extracted with ethyl acetate (20 mL x 2). The combined organic layers were dried over anhydrous sodium sulfate and concentrated under reduced pressure. The residue was applied onto a silica gel column eluting with ethyl acetate/petroleum ether (1:0). This yielded 3.1 g (84%) of benzyl 9-(3-amino-6-chloropyridazin-4-yl)-1-oxa-4,9-diazaspiro[5.5]undecane-4-carboxylate as a yellow solid.
工程4

30mLの密封チューブに、ジオキサン(5mL)およびH2O(1mL)中のベンジル9-(3-アミノ-6-クロロピリダジン-4-イル)-1-オキサ-4,9-ジアザスピロ[5.5]ウンデカン-4-カルボキシレート(3.1g、7.4mmol、1.0当量)、[2-(メトキシメトキシ)フェニル]ボロン酸(2.7g、0.1mmol、2.0当量)、Pd(PPh3)4(859.4mg、0.7mmol、0.1当量)、およびK2CO3(3.1g、0.1mmol、3.0当量)を入れた。得られた混合物を、油浴中で、100℃で12時間撹拌し、次いで、酢酸エチル(20mL×2)で抽出した。合わせた有機相を無水硫酸ナトリウムで乾燥させ、減圧下で濃縮した。残渣を、ジクロロメタン/メタノール(10:1)で溶出するシリカゲルカラム上に適用した。これにより、3.2g(83%)のベンジル9-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-1-オキサ-4,9-ジアザスピロ[5.5]ウンデカン-4-カルボキシレートを黄色の固体として得た。 A 30 mL sealed tube was charged with benzyl 9-(3-amino-6-chloropyridazin-4-yl)-1-oxa-4,9-diazaspiro[5.5]undecane-4-carboxylate (3.1 g, 7.4 mmol, 1.0 equiv.), [2-(methoxymethoxy)phenyl]boronic acid (2.7 g, 0.1 mmol, 2.0 equiv.), Pd(PPh3)4 (859.4 mg, 0.7 mmol, 0.1 equiv.), and K2CO3 (3.1 g, 0.1 mmol, 3.0 equiv.) in dioxane (5 mL) and H2O (1 mL). The resulting mixture was stirred in an oil bath at 100 °C for 12 h and then extracted with ethyl acetate (20 mL x 2). The combined organic phase was dried over anhydrous sodium sulfate and concentrated under reduced pressure. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (10:1). This yielded 3.2 g (83%) of benzyl 9-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-1-oxa-4,9-diazaspiro[5.5]undecane-4-carboxylate as a yellow solid.
工程5

100mLの丸底フラスコで、メタノール(10mL)中のベンジル9-[3-アミノ-6-[2-(メトキシメトキシ)フェニル]ピリダジン-4-イル]-1-オキサ-4,9-ジアザスピロ[5.5]ウンデカン-4-カルボキシレート(3.2g、6.2mmol、1.0当量)の溶液に、Pd(OH)2/C(10%、500mg)を窒素雰囲気下で添加した。次いで、フラスコを真空化し、水素でフラッシュした。反応混合物を、水素バルーンを使用して水素雰囲気下で、室温で水素化し、次いで、セライトパッドを通して濾過し、減圧下で濃縮した。これにより、1.8g(76%)の6-[2-(メトキシメトキシ)フェニル]-4-[1-オキサ-4、9-ジアザスピロ[5.5]ウンデカン-9-イル]ピリダジン-3-アミンを黄色の固体として得た。 In a 100 mL round bottom flask, to a solution of benzyl 9-[3-amino-6-[2-(methoxymethoxy)phenyl]pyridazin-4-yl]-1-oxa-4,9-diazaspiro[5.5]undecane-4-carboxylate (3.2 g, 6.2 mmol, 1.0 equiv.) in methanol (10 mL) was added Pd(OH)2/C (10%, 500 mg) under nitrogen atmosphere. The flask was then evacuated and flushed with hydrogen. The reaction mixture was hydrogenated at room temperature under a hydrogen atmosphere using a hydrogen balloon, then filtered through a celite pad and concentrated under reduced pressure. This gave 1.8 g (76%) of 6-[2-(methoxymethoxy)phenyl]-4-[1-oxa-4,9-diazaspiro[5.5]undecane-9-yl]pyridazin-3-amine as a yellow solid.
工程6

250mLの丸底フラスコに、ジクロロメタン(80mL)中のブタ-2-イン-1,4-ジオール(2.58g、30.00mmol、1.00当量)、酸化銀(10.35g、45.00mmol、1.50当量)、およびヨウ化カリウム(498mg、3.00mmol、0.10当量)の溶液を入れた。次いで、4-メチルベンゼンスルホニルクロリド(5.70g、30.00mmol、1.00当量)を添加した。反応混合物を室温で一晩撹拌した。不溶性固形分を濾過し、濾液を濃縮した。残渣を、酢酸エチル/石油エーテル(1:2)で溶出するシリカゲルカラム上に適用した。これにより、6.13g(85%)の4-ヒドロキシブタ-2-イン-1-イル4-メチルベンゼンスルホネートを黄色の油として得た。 A 250 mL round bottom flask was charged with a solution of but-2-yn-1,4-diol (2.58 g, 30.00 mmol, 1.00 equiv.), silver oxide (10.35 g, 45.00 mmol, 1.50 equiv.), and potassium iodide (498 mg, 3.00 mmol, 0.10 equiv.) in dichloromethane (80 mL). 4-Methylbenzenesulfonyl chloride (5.70 g, 30.00 mmol, 1.00 equiv.) was then added. The reaction mixture was stirred overnight at room temperature. The insoluble solids were filtered and the filtrate was concentrated. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (1:2). This gave 6.13 g (85%) of 4-hydroxybut-2-yn-1-yl 4-methylbenzenesulfonate as a yellow oil.
工程7

250mLの丸底フラスコに、DMF(40mL)中の4-ヒドロキシブタ-2-イン-1-イル4-メチルベンゼンスルホネート(6.13g、25.54mmol、1.10当量)、ベンジルピペラジン-1-カルボキシレート(5.11g、23.22mmol、1.00当量)、および炭酸カリウム(12.82g、92.88mmol、4.00当量)の溶液を入れた。結果として生じる混合物を室温で一晩撹拌した。次いで、反応混合物を水(400mL)によってクエンチし、酢酸エチル(200mL×2)で抽出した。合わせた有機相を硫酸ナトリウムで乾燥させ、濃縮した。残渣を、酢酸エチル/石油エーテル(1:1)で溶出するシリカゲルカラム上に適用した。これにより、2.01g(30%)のベンジル4-(4-ヒドロキシブタ-2-イン-1-イル)ピペラジン-1-カルボキシレートを黄色の油として得た。 A 250 mL round bottom flask was charged with a solution of 4-hydroxybut-2-yn-1-yl 4-methylbenzenesulfonate (6.13 g, 25.54 mmol, 1.10 equiv.), benzyl piperazine-1-carboxylate (5.11 g, 23.22 mmol, 1.00 equiv.), and potassium carbonate (12.82 g, 92.88 mmol, 4.00 equiv.) in DMF (40 mL). The resulting mixture was stirred overnight at room temperature. The reaction mixture was then quenched with water (400 mL) and extracted with ethyl acetate (200 mL x 2). The combined organic phases were dried over sodium sulfate and concentrated. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (1:1). This gave 2.01 g (30%) of benzyl 4-(4-hydroxybut-2-yn-1-yl)piperazine-1-carboxylate as a yellow oil.
工程8

250mLの丸底フラスコ、ジエチルエーテル(100mL)中のベンジル4-(4-ヒドロキシブタ-2-イン-1-イル)ピペラジン-1-カルボキシレート(2.01g、6.98mmol、1.00当量)および水酸化カリウム(3.91g、69.80mmol、10.00当量)の溶液を入れた。次いで、4-メチルベンゼンスルホニルクロリド(1.99g、10.47mmol、1.50当量)を添加した。反応混合物を室温で一晩撹拌した。不溶性固形分を濾過した。濾液を水(100mL)で洗浄し、無水硫酸ナトリウムで乾燥させ、真空下で濃縮した。これにより、1.80g(59%)のベンジル4-(4-(トシルオキシ)ブタ-2-イン-1-イル)ピペラジン-1-カルボキシレートを黄色の油として得た。 A 250 mL round bottom flask was charged with a solution of benzyl 4-(4-hydroxybut-2-yn-1-yl)piperazine-1-carboxylate (2.01 g, 6.98 mmol, 1.00 equiv.) and potassium hydroxide (3.91 g, 69.80 mmol, 10.00 equiv.) in diethyl ether (100 mL). 4-Methylbenzenesulfonyl chloride (1.99 g, 10.47 mmol, 1.50 equiv.) was then added. The reaction mixture was stirred overnight at room temperature. The insoluble solids were filtered off. The filtrate was washed with water (100 mL), dried over anhydrous sodium sulfate, and concentrated under vacuum. This afforded 1.80 g (59%) of benzyl 4-(4-(tosyloxy)but-2-yn-1-yl)piperazine-1-carboxylate as a yellow oil.
工程9

250mLの丸底フラスコに、アセトニトリル(70mL)中のベンジル4-(4-(トシルオキシ)ブタ-2-イン-1-イル)ピペラジン-1-カルボキシレート(1.80g、4.07mmol、2.50当量)、6-(2-(メトキシメトキシ)フェニル)-4-(1-オキサ-4,9-ジアザスピロ[5.5]ウンデカン-9-イル)ピリダジン-3-アミン(627mg、1.63mmol、1.00当量)、ヨウ化ナトリウム(366mg、2.44mmol、1.50当量)、炭酸カリウム(900mg、6.52mmol、4.0当量)の溶液を入れた。得られた混合物を70℃で一晩撹拌した。反応混合物を水(100mL)で希釈し、酢酸エチル(200mL×2)で抽出した。合わせた有機相を濃縮し、残渣を、メタノール/ジクロロメタン(1:10)で溶出するシリカゲルカラム上に適用した。これにより、180mg(17%)のベンジル4-(4-(9-(3-アミノ-6-(2-(メトキシメトキシ)フェニル)ピリダジン-4-イル)-1-オキサ-4,9-ジアザスピロ[5.5]ウンデカン-4-イル)ブタ-2-イン-1-イル)ピペラジン-1-カルボキシレートを茶色の固体として得た。 A 250 mL round bottom flask was charged with a solution of benzyl 4-(4-(tosyloxy)but-2-yn-1-yl)piperazine-1-carboxylate (1.80 g, 4.07 mmol, 2.50 equiv.), 6-(2-(methoxymethoxy)phenyl)-4-(1-oxa-4,9-diazaspiro[5.5]undecan-9-yl)pyridazin-3-amine (627 mg, 1.63 mmol, 1.00 equiv.), sodium iodide (366 mg, 2.44 mmol, 1.50 equiv.), potassium carbonate (900 mg, 6.52 mmol, 4.0 equiv.) in acetonitrile (70 mL). The resulting mixture was stirred at 70 °C overnight. The reaction mixture was diluted with water (100 mL) and extracted with ethyl acetate (200 mL × 2). The combined organic phases were concentrated and the residue was applied onto a silica gel column eluted with methanol/dichloromethane (1:10). This gave 180 mg (17%) of benzyl 4-(4-(9-(3-amino-6-(2-(methoxymethoxy)phenyl)pyridazin-4-yl)-1-oxa-4,9-diazaspiro[5.5]undecan-4-yl)but-2-yn-1-yl)piperazine-1-carboxylate as a brown solid.
工程10

100のmL丸底フラスコで、メタノール(40mL)中のベンジル4-(4-(9-(3-アミノ-6-(2-(メトキシメトキシ)フェニル)ピリダジン-4-イル)-1-オキサ-4,9-ジアザスピロ[5.5]ウンデカン-4-イル)ブタ-2-イン-1-イル)ピペラジン-1-カルボキシレート(1.60g、2.44mmol、1.00当量)の溶液に、二酸化白金(221.8mg、0.98mmol、0.40当量)を窒素雰囲気下で添加した。フラスコを真空化し、水素で流した。反応混合物を、水素バルーンを使用して室温で4時間水素化した。混合物を、セライトパッドを通して濾過し、濾液を減圧下で濃縮した。残渣を、メタノール/ジクロロメタン(1:10)で溶出するシリカゲルカラム上に適用した。これにより、444.0mg(28%)のベンジル4-(4-(9-(3-アミノ-6-(2-(メトキシメトキシ)フェニル)ピリダジン-4-イル)-1-オキサ-4,9-ジアザスピロ[5.5]ウンデカン-4-イル)ブチル)ピペラジン-1-カルボキシレートを茶色の固体として得た。 To a solution of benzyl 4-(4-(9-(3-amino-6-(2-(methoxymethoxy)phenyl)pyridazin-4-yl)-1-oxa-4,9-diazaspiro[5.5]undecan-4-yl)but-2-yn-1-yl)piperazine-1-carboxylate (1.60 g, 2.44 mmol, 1.00 equiv.) in methanol (40 mL) in a 100 mL round bottom flask, platinum dioxide (221.8 mg, 0.98 mmol, 0.40 equiv.) was added under nitrogen atmosphere. The flask was evacuated and flushed with hydrogen. The reaction mixture was hydrogenated at room temperature using a hydrogen balloon for 4 h. The mixture was filtered through a celite pad and the filtrate was concentrated under reduced pressure. The residue was applied onto a silica gel column eluting with methanol/dichloromethane (1:10). This yielded 444.0 mg (28%) of benzyl 4-(4-(9-(3-amino-6-(2-(methoxymethoxy)phenyl)pyridazin-4-yl)-1-oxa-4,9-diazaspiro[5.5]undecan-4-yl)butyl)piperazine-1-carboxylate as a brown solid.
工程11

100mLの丸底フラスコに、THF(8mL)中のベンジル4-(4-(9-(3-アミノ-6-(2-(メトキシメトキシ)フェニル)ピリダジン-4-イル)-1-オキサ-4,9-ジアザスピロ[5.5]ウンデカン-4-イル)ブチル)ピペラジン-1-カルボキシレート(444mg、0.67mmol、1.00当量)の溶液を入れ、それにTHF(1.0M、8mL)中の水素化トリエチルホウ素リチウムの溶液を添加した。得られた溶液を室温で3時間撹拌した。反応混合物を、飽和NH4Cl溶液(30mL)によってクエンチし、ジクロロメタン(30mL×2)で抽出した。合わせた有機相を硫酸ナトリウムで乾燥させ、減圧下で濃縮した。これにより、292mg(83%)の6-(2-(メトキシメトキシ)フェニル)-4-(4-(4-(ピペラジン-1-イル)ブチル)-1-オキサ-4,9-ジアザスピロ[5.5]ウンデカン-9-イル)ピリダジン-3-アミンを茶色の油として得た。 A 100 mL round bottom flask was charged with a solution of benzyl 4-(4-(9-(3-amino-6-(2-(methoxymethoxy)phenyl)pyridazin-4-yl)-1-oxa-4,9-diazaspiro[5.5]undecan-4-yl)butyl)piperazine-1-carboxylate (444 mg, 0.67 mmol, 1.00 equiv.) in THF (8 mL), to which was added a solution of lithium triethylborohydride in THF (1.0 M, 8 mL). The resulting solution was stirred at room temperature for 3 h. The reaction mixture was quenched by saturated NH4Cl solution (30 mL) and extracted with dichloromethane (30 mL x 2). The combined organic phase was dried over sodium sulfate and concentrated under reduced pressure. This gave 292 mg (83%) of 6-(2-(methoxymethoxy)phenyl)-4-(4-(4-(piperazin-1-yl)butyl)-1-oxa-4,9-diazaspiro[5.5]undecan-9-yl)pyridazin-3-amine as a brown oil.
他の実施例について上に記載されるものと類似の手順を使用して、以下のスキームに従って、6-(2-(メトキシメトキシ)フェニル)-4-(4-(4-(ピペラジン-1-イル)ブチル)-1-オキサ-4,9-ジアザスピロ[5.5]ウンデカン-9-イル)ピリダジン-3-アミンを最終化合物に変換した。
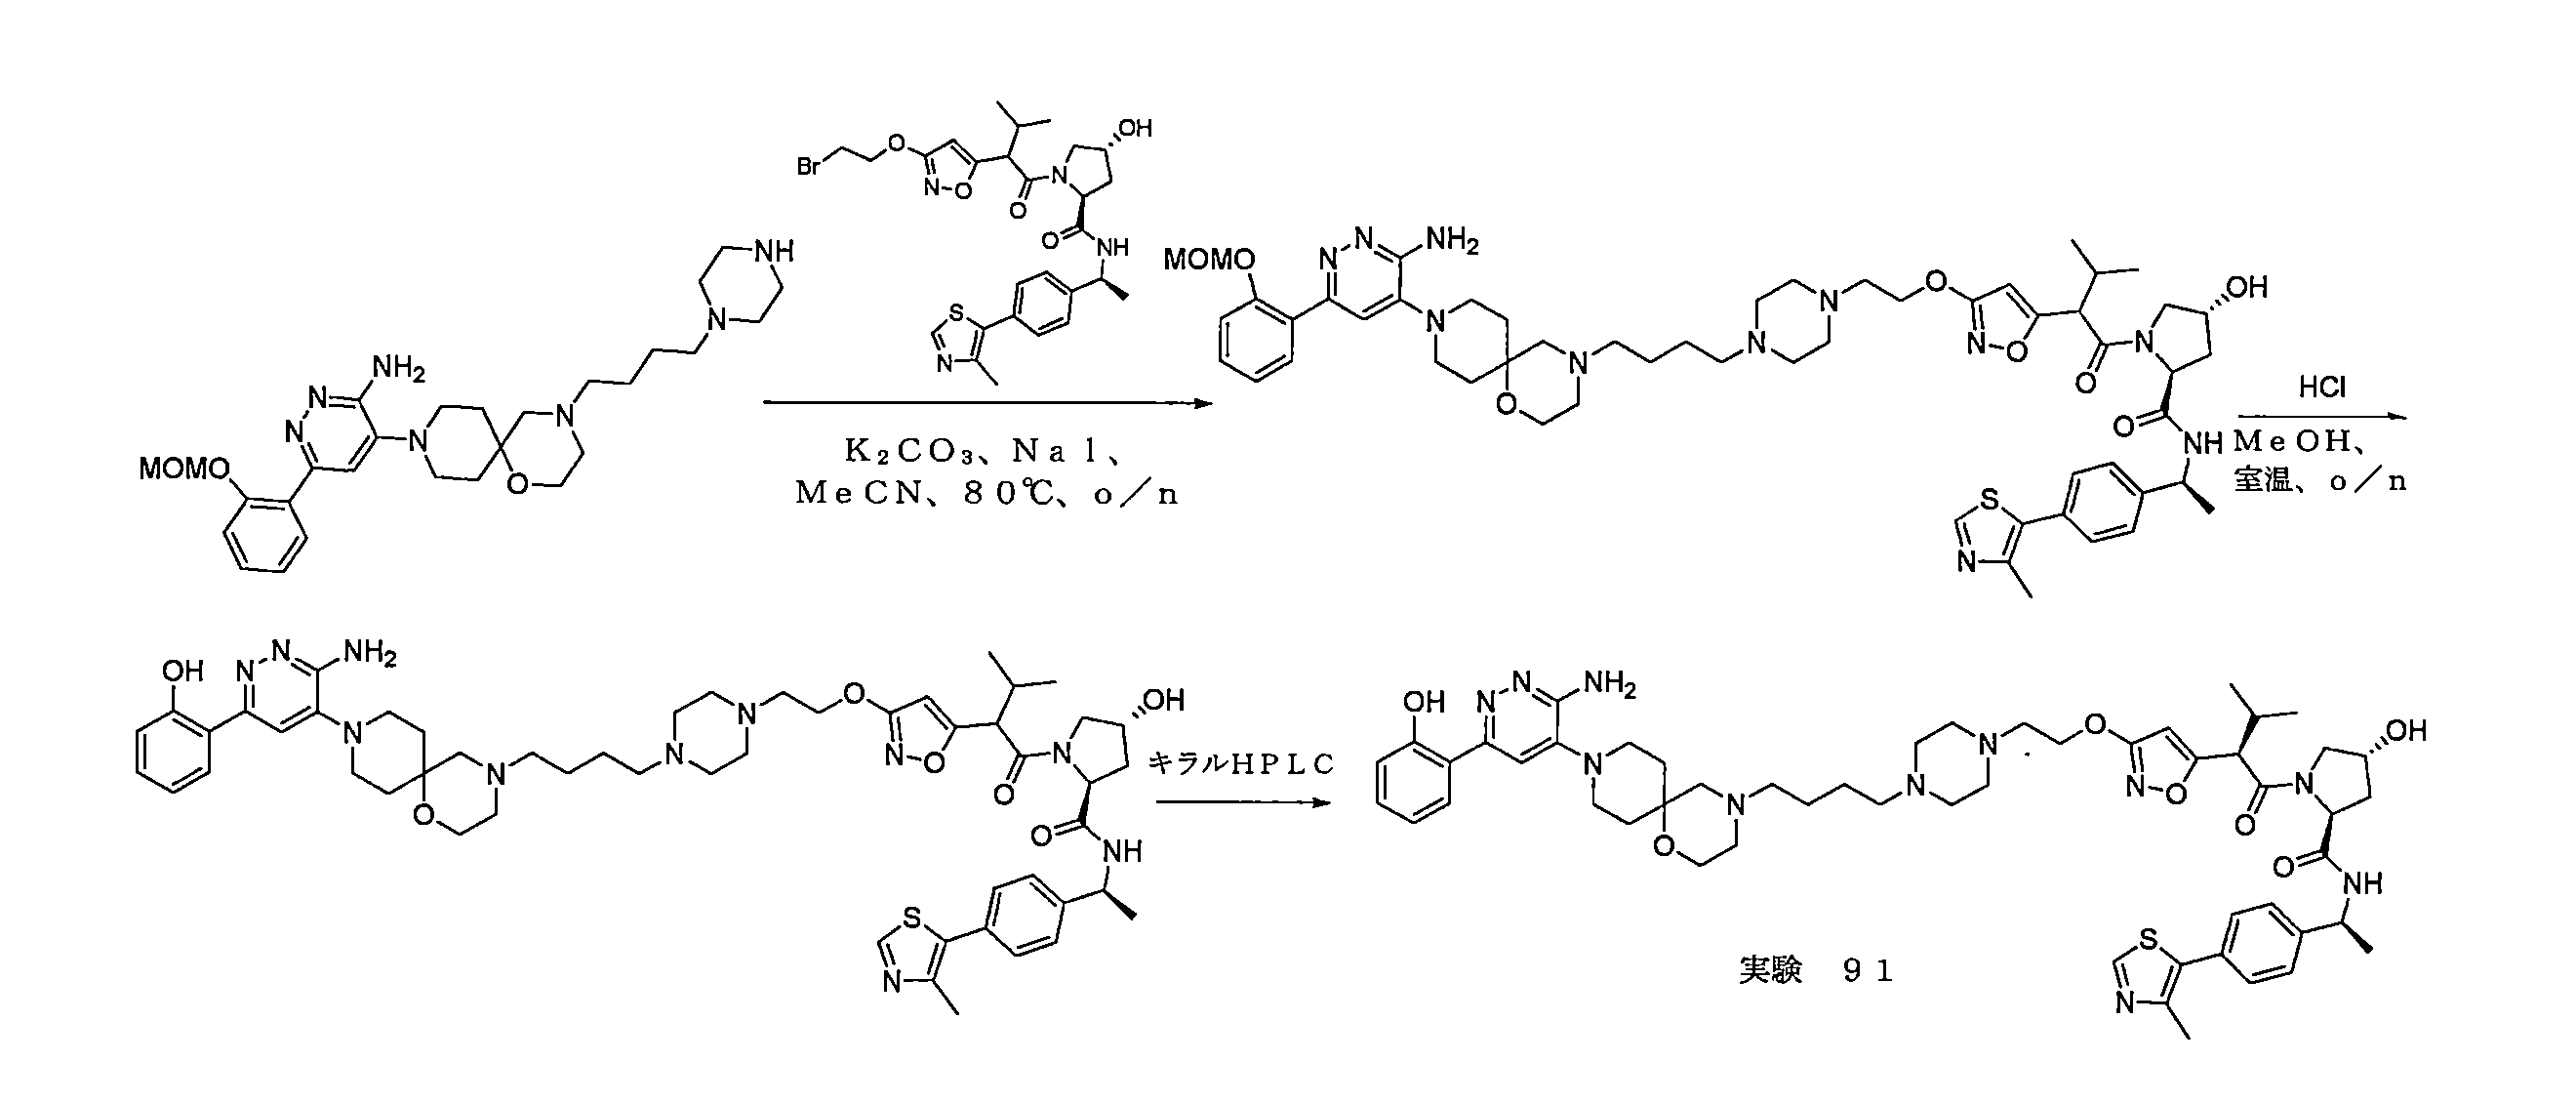
例示的な化合物205を、類似の手順を使用して調製した。 Exemplary compound 205 was prepared using a similar procedure.
例示的な化合物188の例示的な合成
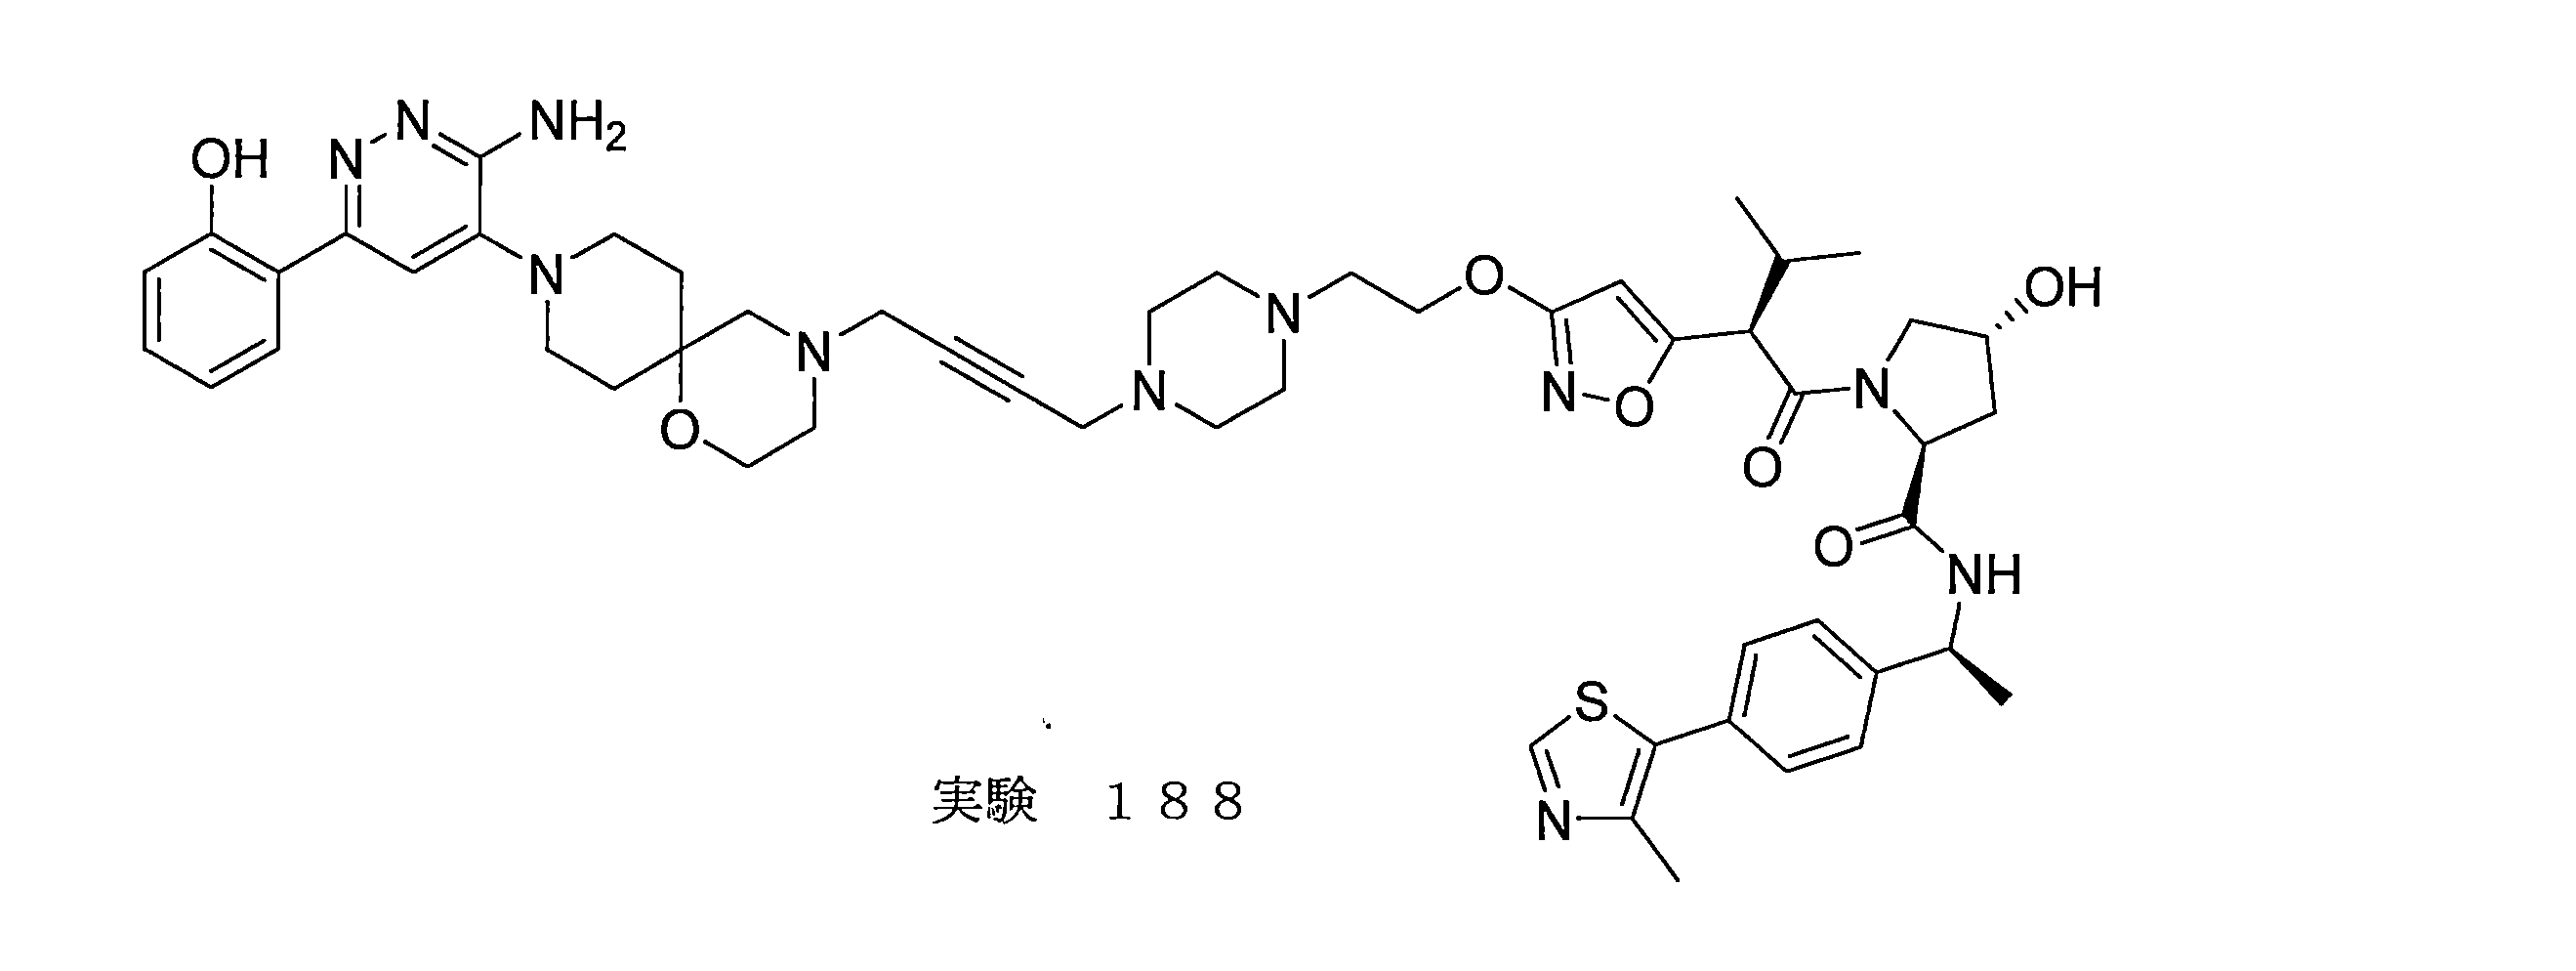
例示的化合物91について上に記載されるものと類似の手順を使用して、以下のスキームに従って調製した。

例示的な化合物187を、類似の手順を使用して調製した。 Exemplary compound 187 was prepared using a similar procedure.
例示的な化合物192の例示的な合成

他の実施例について記載される手順、ならびに当業者に明らかである手順を使用して、以下のスキームに従って、ベンジル4-(2-((1r,3r)-3-(2-(9-(3-アミノ-6-(2-(メトキシメトキシ)フェニル)ピリダジン-4-イル)-1-オキサ-4,9-ジアザスピロ[5.5]ウンデカン-4-イル)エトキシ)シクロブトキシ)エチル)ピペラジン-1-カルボキシレーを調製した。

例示的な化合物91について記載されるものと類似の手順を使用して、ベンジル4-(2-((1r,3r)-3-(2-(9-(3-アミノ-6-(2-(メトキシメトキシ)フェニル)ピリダジン-4-イル)-1-オキサ-4,9-ジアザスピロ[5.5]ウンデカン-4-イル)エトキシ)シクロブトキシ)エチル)ピペラジン-1-カルボキシレートを最終化合物に変換した。 Using a procedure similar to that described for exemplary compound 91, benzyl 4-(2-((1r,3r)-3-(2-(9-(3-amino-6-(2-(methoxymethoxy)phenyl)pyridazin-4-yl)-1-oxa-4,9-diazaspiro[5.5]undecan-4-yl)ethoxy)cyclobutoxy)ethyl)piperazine-1-carboxylate was converted to the final compound.
例示的な化合物212および213の例示的な合成
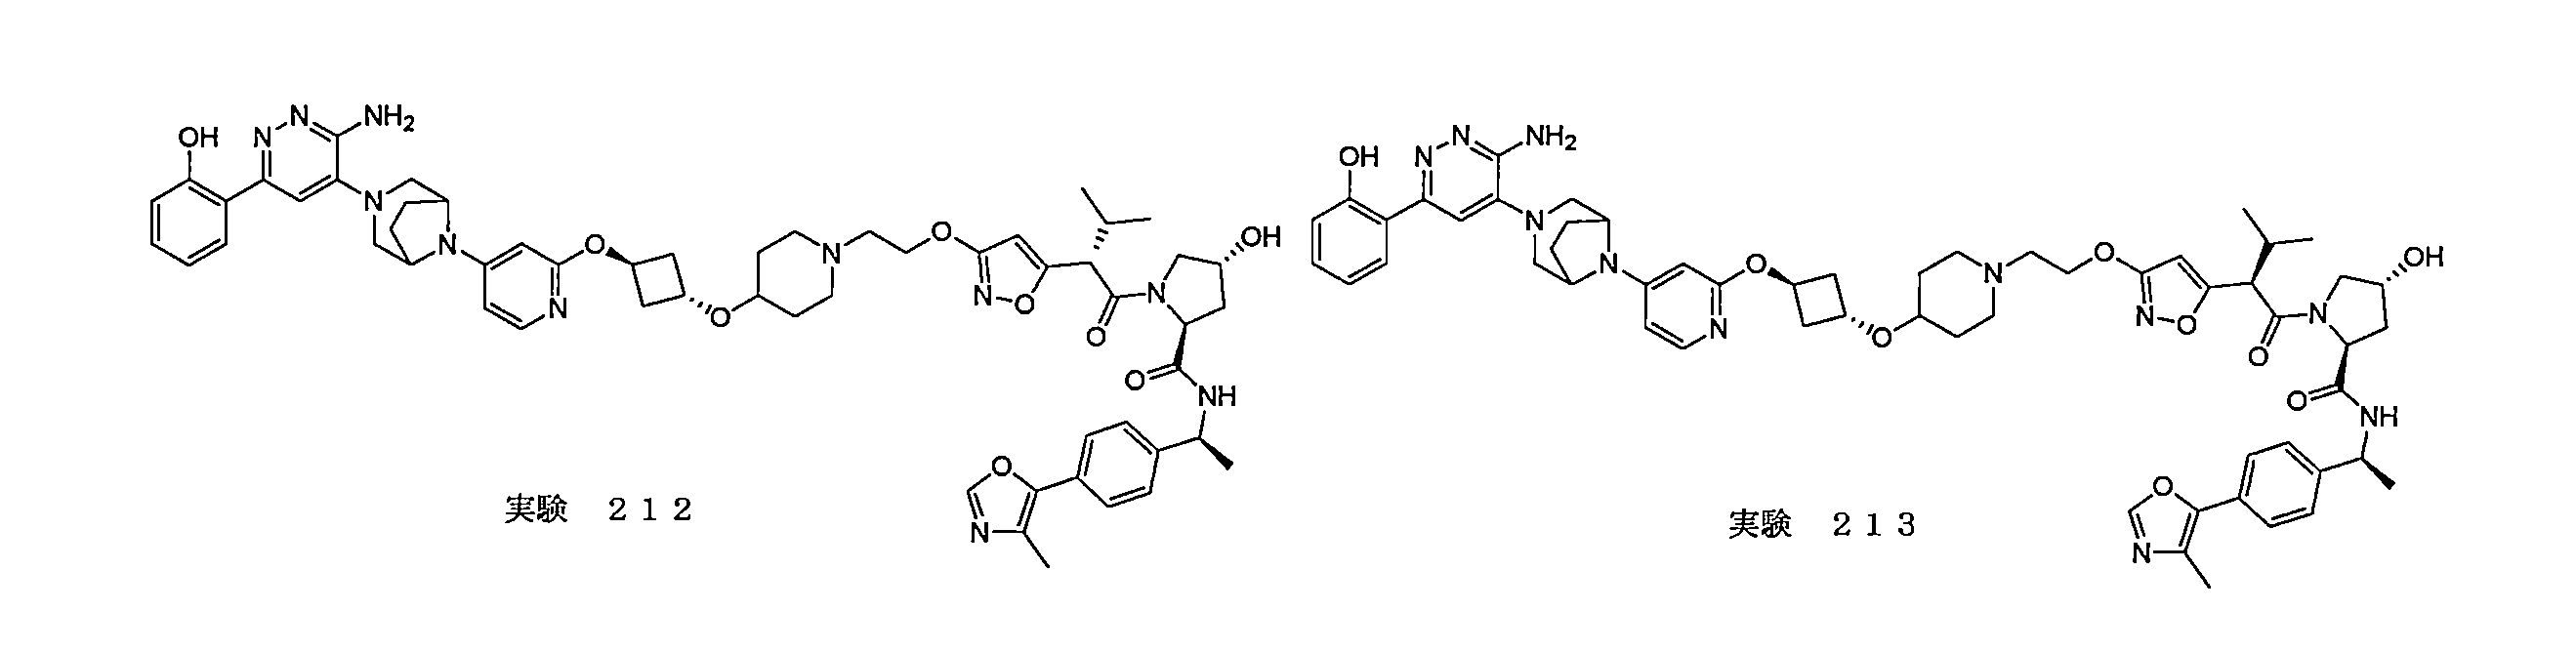
例示的な化合物117の工程1および2、続いて例示的な化合物180について記載される工程に記載されるものと類似の手順を使用して調製した。 Prepared using procedures similar to those described in steps 1 and 2 of exemplary compound 117, followed by the steps described for exemplary compound 180.
例示的な化合物214および215の例示的な合成
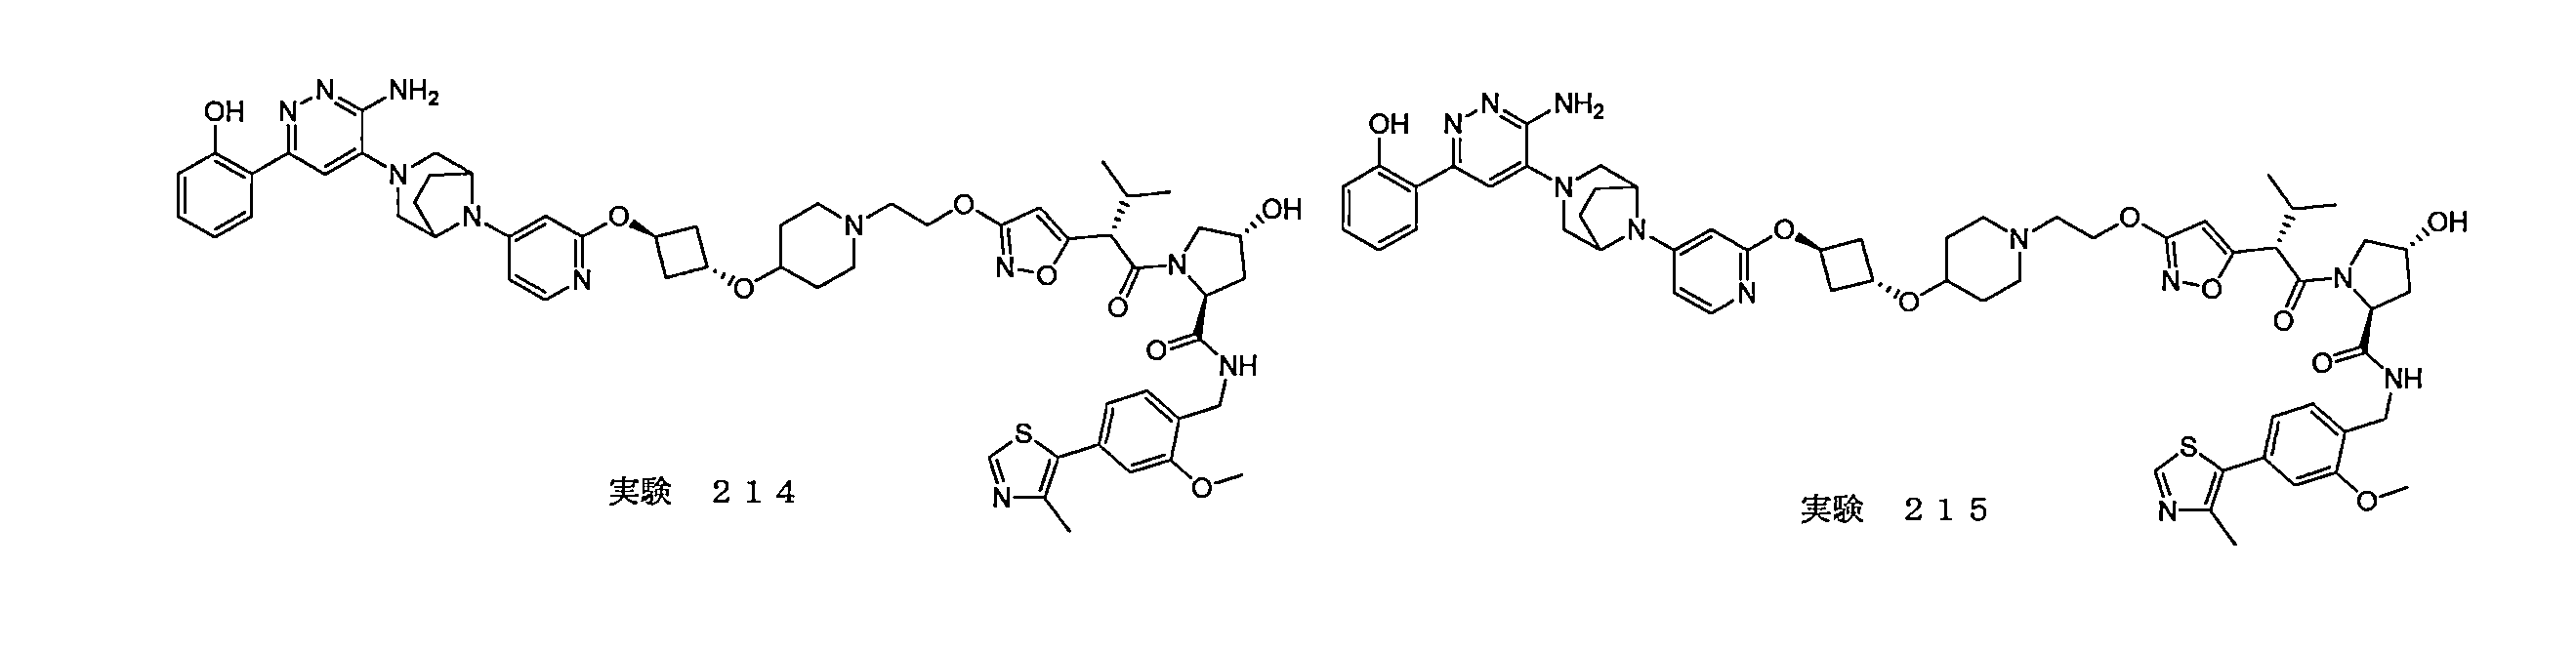
当業者に明らかである標準的な手順を使用して、以下のスキームに従って、tert-ブチル(2S,4R)-4-ヒドロキシ-2-((2-メトキシ-4-(4-メチルチアゾール-5-イル)ベンジル)カルバモイル)ピロリジン-1-カルボキシレートを調製した。
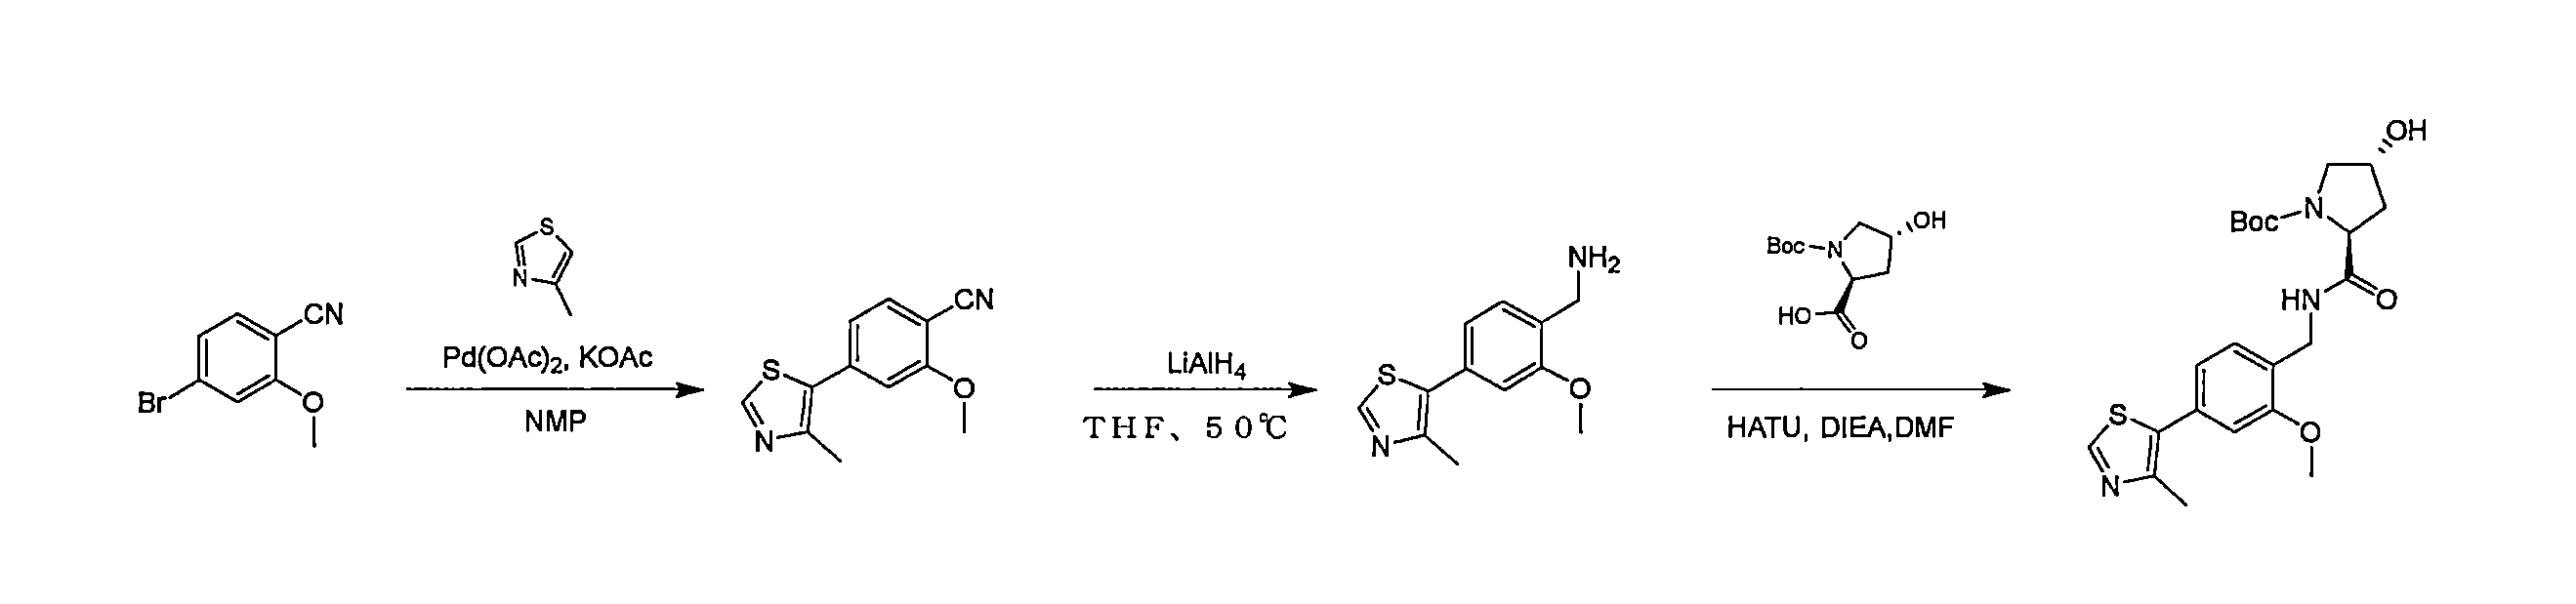
上記の他の実施例について記載される標準的な手順を使用して、tert-ブチル(2S,4R)-4-ヒドロキシ-2-((2-メトキシ-4-(4-メチルチアゾール-5-イル)ベンジル)カルバモイル)ピロリジン-1-カルボキシレートを最終化合物に変換した。 tert-Butyl (2S,4R)-4-hydroxy-2-((2-methoxy-4-(4-methylthiazol-5-yl)benzyl)carbamoyl)pyrrolidine-1-carboxylate was converted to the final compound using standard procedures as described for other examples above.
例示的な化合物238の例示的な合成

工程1
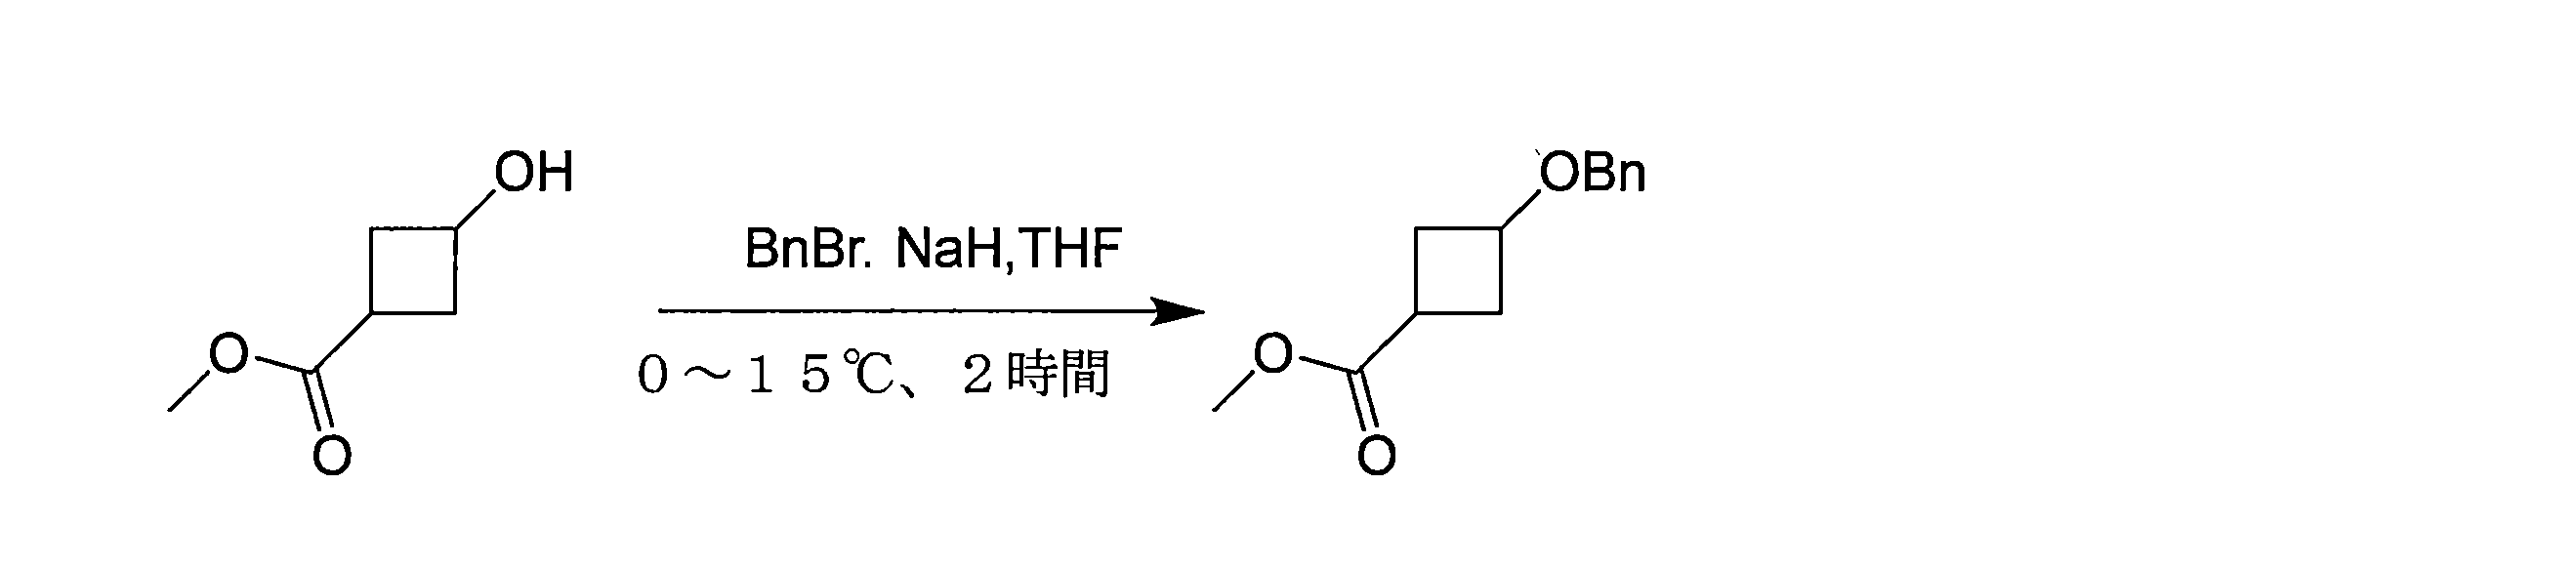
DMF(150mL)中のメチル3-ヒドロキシシクロブタンカルボキシレート(14.2g、109.11mmol、1.00当量)の溶液に、水素化ナトリウム(5.24g、130.93mmol、純度60%、1.20当量)を添加し、混合物を0℃で0.5時間撹拌した。その後、臭化ベンジル(20.53g、120.02mmol、14.26mL、1.1当量)を添加し、混合物を25℃で2時間撹拌した。反応混合物を、25℃で、飽和塩化アンモニウム溶液(200mL)によってクエンチし、次いで、水(100mL)で希釈し、酢酸エチル(80mL×3)で抽出した。合わせた有機層をブライン(150mL×2)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、減圧下で濃縮して、残渣を得た。残渣を、フラッシュシリカゲルクロマトグラフィー(0~20%酢酸エチル/石油エーテル勾配)によって精製した。化合物メチル3-ベンジルオキシシクロブタンカルボキシレート(8.9g、40.41mmol、収率37%)を淡い黄色の油として得た。 To a solution of methyl 3-hydroxycyclobutanecarboxylate (14.2 g, 109.11 mmol, 1.00 equiv.) in DMF (150 mL), sodium hydride (5.24 g, 130.93 mmol, 60% purity, 1.20 equiv.) was added and the mixture was stirred at 0° C. for 0.5 h. Then, benzyl bromide (20.53 g, 120.02 mmol, 14.26 mL, 1.1 equiv.) was added and the mixture was stirred at 25° C. for 2 h. The reaction mixture was quenched by saturated ammonium chloride solution (200 mL) at 25° C., then diluted with water (100 mL) and extracted with ethyl acetate (80 mL×3). The combined organic layers were washed with brine (150 mL×2), dried over anhydrous sodium sulfate, filtered and concentrated under reduced pressure to give a residue. The residue was purified by flash silica gel chromatography (0-20% ethyl acetate/petroleum ether gradient). The compound methyl 3-benzyloxycyclobutanecarboxylate (8.9 g, 40.41 mmol, 37% yield) was obtained as a pale yellow oil.
工程2
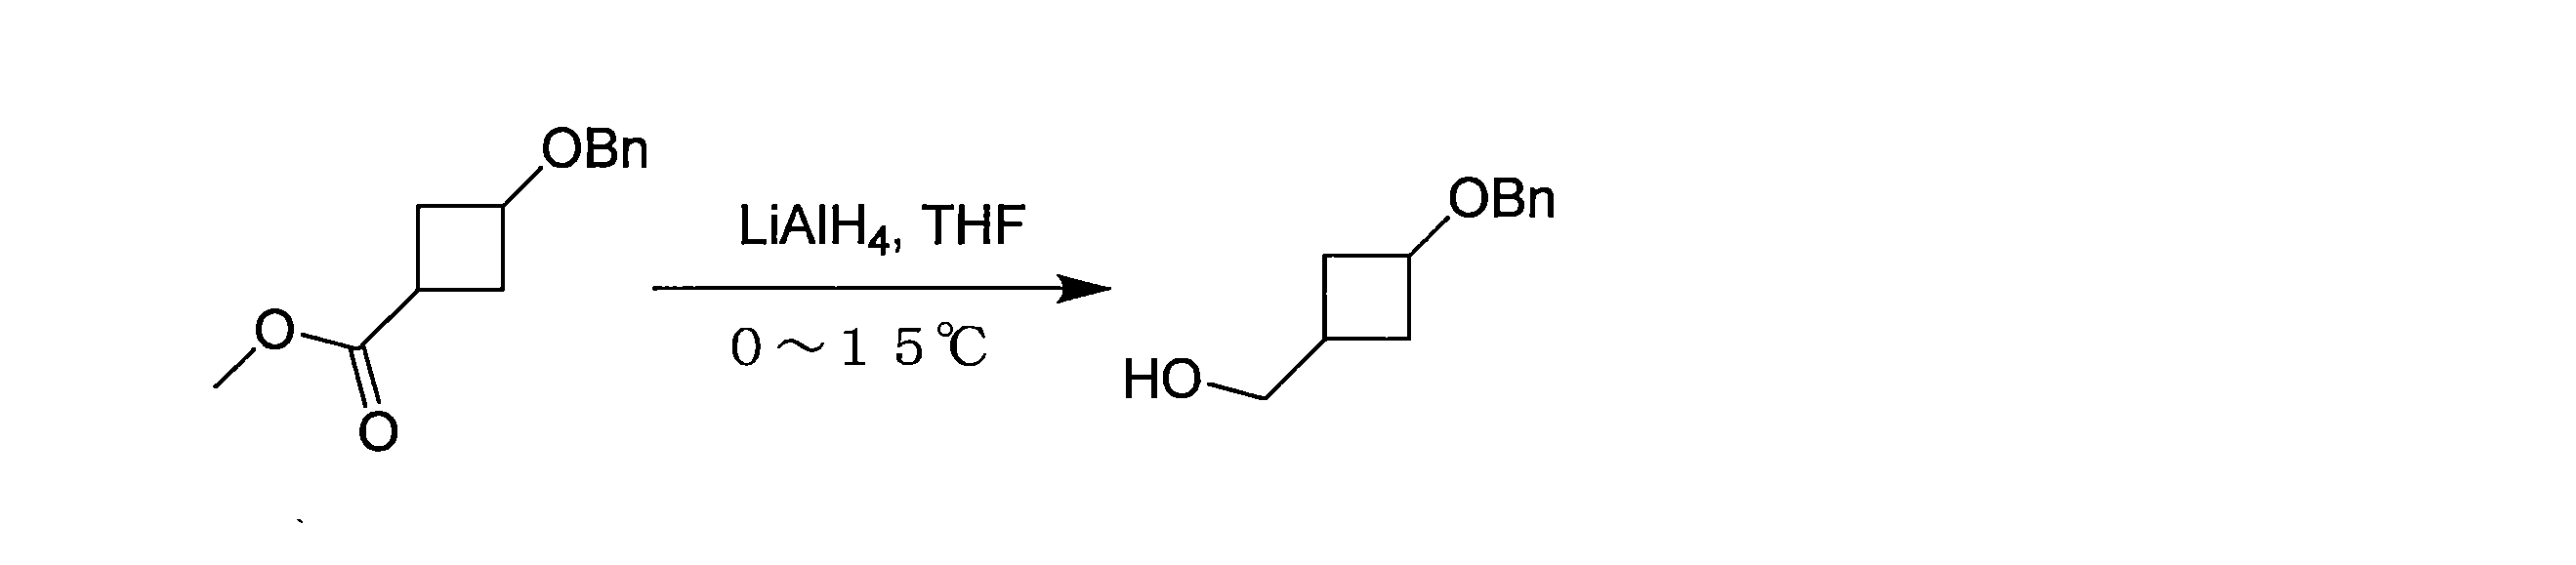
テトラヒドロフラン(80mL)中の水素化リチウムアルミニウム(1.53g、40.41mmol、1.00当量)の溶液に、メチル3-ベンジルオキシシクロブタンカルボキシレート(8.90g、40.41mmol、1.00当量)を0℃で添加し、混合物を25℃で2時間撹拌した。反応混合物を、25℃で、水(1mL)、水酸化ナトリウム溶液(15%、2mL)、および水(3mL)によってクエンチし、次いで、酢酸エチル(80mL)で希釈し、酢酸エチル(15mL×3)で抽出した。合わせた有機層をブライン(50mL×2)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、減圧下で濃縮して、残渣を得た。残渣を、フラッシュシリカゲルクロマトグラフィー(0~50%酢酸エチル/石油エーテル)によって精製した。化合物(3-ベンジルオキシシクロブチル)メタノール(7.4g、38.5mmol、収率95%)を無色の油として得た。 To a solution of lithium aluminum hydride (1.53 g, 40.41 mmol, 1.00 equiv.) in tetrahydrofuran (80 mL) was added methyl 3-benzyloxycyclobutanecarboxylate (8.90 g, 40.41 mmol, 1.00 equiv.) at 0° C., and the mixture was stirred at 25° C. for 2 h. The reaction mixture was quenched with water (1 mL), sodium hydroxide solution (15%, 2 mL), and water (3 mL) at 25° C., then diluted with ethyl acetate (80 mL) and extracted with ethyl acetate (15 mL×3). The combined organic layers were washed with brine (50 mL×2), dried over anhydrous sodium sulfate, filtered, and concentrated under reduced pressure to give a residue. The residue was purified by flash silica gel chromatography (0-50% ethyl acetate/petroleum ether). Compound (3-benzyloxycyclobutyl)methanol (7.4 g, 38.5 mmol, 95% yield) was obtained as a colorless oil.
工程3

ジクロロメタン(125mL)中の(3-ベンジルオキシシクロブチル)メタノール(7.4g、38.49mmol、1.00当量)および3,4-ジヒドロ-2H-ピラン(16.19g、192.5mmol、17.6mL、5.00当量)の溶液に、p-トルエンスルホン酸ピリジニウム(967mg、3.85mmol、0.10当量)を添加し、混合物を25℃で12時間撹拌した。水(200mL)を混合物に添加し、得られた混合物をジクロロメタン(100mL×2)で抽出した。次いで、合わせた有機層をブライン(150mL×2)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、減圧下で濃縮して、残渣を得た。残渣を、カラムクロマトグラフィー(SiO2、石油エーテル:酢酸エチル=1:0~20:1)によって精製した。化合物2-[(3-ベンジルオキシシクロブチル)メトキシ]テトラヒドロピラン(10.20g、36.9mmol、収率95%)を無色の油として得た。 To a solution of (3-benzyloxycyclobutyl)methanol (7.4 g, 38.49 mmol, 1.00 equiv.) and 3,4-dihydro-2H-pyran (16.19 g, 192.5 mmol, 17.6 mL, 5.00 equiv.) in dichloromethane (125 mL), pyridinium p-toluenesulfonate (967 mg, 3.85 mmol, 0.10 equiv.) was added and the mixture was stirred at 25 °C for 12 h. Water (200 mL) was added to the mixture and the resulting mixture was extracted with dichloromethane (100 mL x 2). The combined organic layers were then washed with brine (150 mL x 2), dried over anhydrous sodium sulfate, filtered, and concentrated under reduced pressure to give a residue. The residue was purified by column chromatography (SiO2, petroleum ether: ethyl acetate = 1: 0 to 20: 1). The compound 2-[(3-benzyloxycyclobutyl)methoxy]tetrahydropyran (10.20 g, 36.9 mmol, 95% yield) was obtained as a colorless oil.
工程4
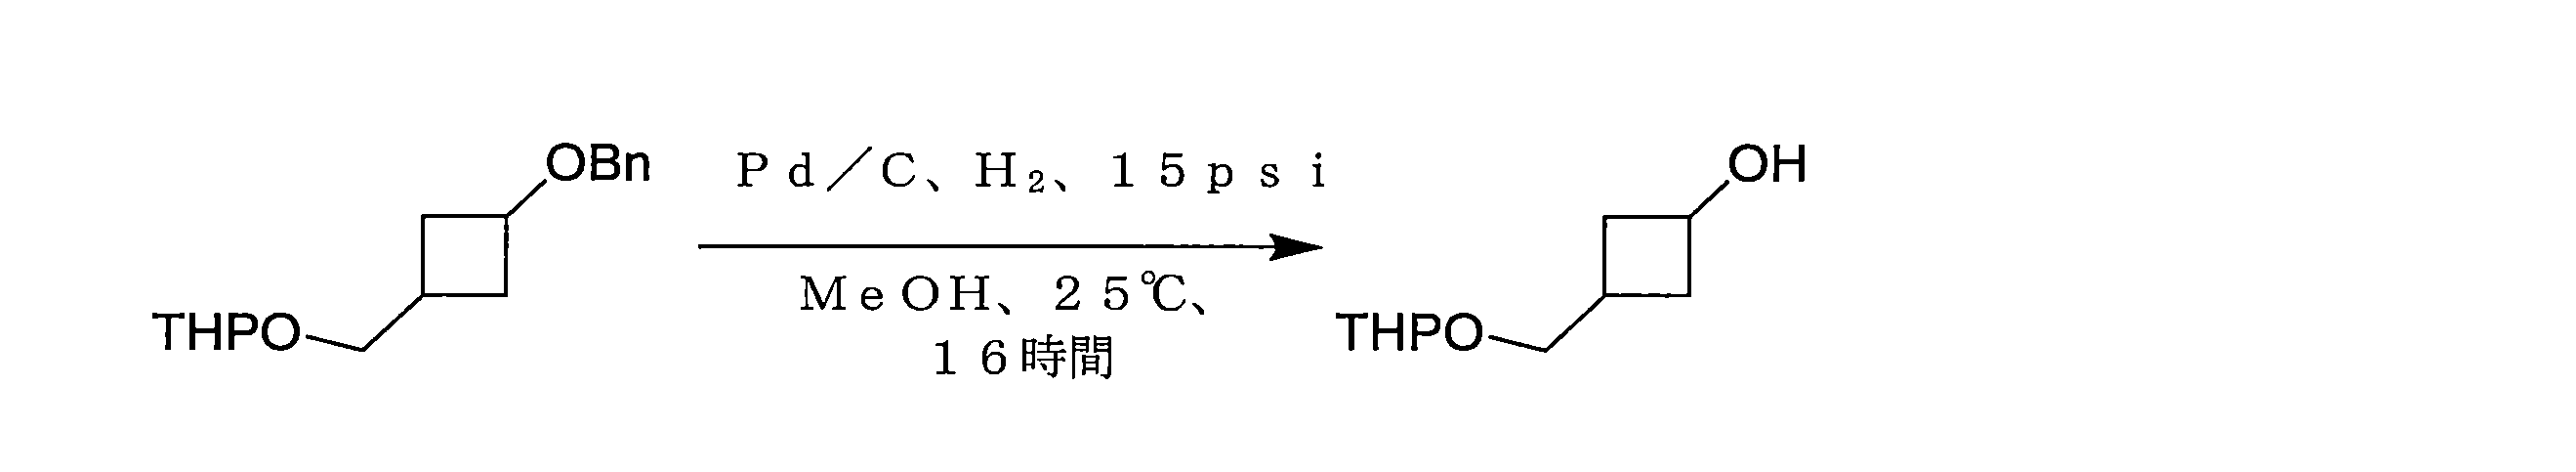
メタノール(100mL)中の2-[(3-ベンジルオキシシクロブチル)メトキシ]テトラヒドロピラン(10.20g、36.9mmol、1.00当量)の溶液に、10%パラジウム炭素(2g、36.91mmol)を窒素下で添加した。懸濁液を真空下で脱気し、水素で数回パージした。混合物を、水素(15psi)下で、25℃で16時間撹拌した。反応物を濾過し、減圧下で濃縮して、残渣を得た。化合物3-(テトラヒドロピラン-2-イルオキシメチル)シクロブタノール(6.28g、33.7mmol、収率91%)を無色の油として得た。 To a solution of 2-[(3-benzyloxycyclobutyl)methoxy]tetrahydropyran (10.20 g, 36.9 mmol, 1.00 equiv.) in methanol (100 mL) was added 10% palladium on carbon (2 g, 36.91 mmol) under nitrogen. The suspension was degassed under vacuum and purged with hydrogen several times. The mixture was stirred under hydrogen (15 psi) at 25°C for 16 h. The reaction was filtered and concentrated under reduced pressure to give a residue. Compound 3-(tetrahydropyran-2-yloxymethyl)cyclobutanol (6.28 g, 33.7 mmol, 91% yield) was obtained as a colorless oil.
工程5

トルエン(40mL)中のピリジン-4-オール(4g、42.06mmol、1当量)、3-(テトラヒドロピラン-2-イルオキシメチル)シクロブタノール(8.62g、46.27mmol、1.1当量)、およびトリフェニルホスフィン(33.10g、126.18mmol、3当量)の溶液に、トルエン(20mL)中のジイソプロピルアゾジカルボキシレート(25.52g、126.18mmol、24.5mL、3当量)を窒素下で、0℃で、滴下で添加した。反応混合物を100℃で12時間撹拌した。混合物を真空中で濃縮して、トルエンのほとんどを除去した。水(150mL)および酢酸エチル(100mL)を混合物に添加し、塩酸溶液(2M)を混合物に添加して、pHを約3に調整し、得られた混合物を酢酸エチル(100mL×2)で抽出した。飽和重炭酸ナトリウムを水相に添加して、pHを約8に調整し、得られた混合物を酢酸エチル(150mL×5)で抽出した。合わせた有機相をブライン(150mL)で洗浄し、硫酸ナトリウムで乾燥させ、濾過し、真空中で濃縮した。粗生成物を、分取HPLC(カラム:Kromasil Eternity XT 250*80mm*10um;移動相:[水(0.05%水酸化アンモニアv/v)-ACN];B%:5%~30%、31分)によって精製した。((1r,3r)-3-(ピリジン-4-イルオキシ)シクロブチル)メタノール(4.2g、23.0mmol、収率54%)を淡い黄色の油として得た。 To a solution of pyridin-4-ol (4 g, 42.06 mmol, 1 eq.), 3-(tetrahydropyran-2-yloxymethyl)cyclobutanol (8.62 g, 46.27 mmol, 1.1 eq.), and triphenylphosphine (33.10 g, 126.18 mmol, 3 eq.) in toluene (40 mL), diisopropyl azodicarboxylate (25.52 g, 126.18 mmol, 24.5 mL, 3 eq.) in toluene (20 mL) was added dropwise under nitrogen at 0°C. The reaction mixture was stirred at 100°C for 12 hours. The mixture was concentrated in vacuum to remove most of the toluene. Water (150 mL) and ethyl acetate (100 mL) were added to the mixture, hydrochloric acid solution (2 M) was added to the mixture to adjust the pH to about 3, and the resulting mixture was extracted with ethyl acetate (100 mL x 2). Saturated sodium bicarbonate was added to the aqueous phase to adjust the pH to about 8, and the resulting mixture was extracted with ethyl acetate (150 mL x 5). The combined organic phase was washed with brine (150 mL), dried over sodium sulfate, filtered, and concentrated in vacuum. The crude product was purified by preparative HPLC (column: Kromasil Eternity XT 250*80 mm*10 um; mobile phase: [water (0.05% ammonia hydroxide v/v)-ACN]; B%: 5%-30%, 31 min). ((1r,3r)-3-(pyridin-4-yloxy)cyclobutyl)methanol (4.2 g, 23.0 mmol, 54% yield) was obtained as a pale yellow oil.
工程6

トルエン(40mL)中の((1r,3r)-3-(ピリジン-4-イルオキシ)シクロブチル)メタノール(4.2g、23.44mmol、1当量)の溶液に、臭化ベンジル(4.41g、25.78mmol、3.1mL、1.1当量)を添加した。反応混合物を80℃で4時間撹拌した。混合物を真空中で濃縮した。残渣を、石油エーテルおよび酢酸エチル(40mL、V/V=1/1)で粉砕した。1-ベンジル-4-((1r,3r)-3-(ヒドロキシメチル)シクロブトキシ)ピリジン-1-イウム(8g、22.8mmol、収率97%)を白色の固体として得た。 To a solution of ((1r,3r)-3-(pyridin-4-yloxy)cyclobutyl)methanol (4.2 g, 23.44 mmol, 1 equiv.) in toluene (40 mL), benzyl bromide (4.41 g, 25.78 mmol, 3.1 mL, 1.1 equiv.) was added. The reaction mixture was stirred at 80°C for 4 h. The mixture was concentrated in vacuo. The residue was triturated with petroleum ether and ethyl acetate (40 mL, V/V=1/1). 1-benzyl-4-((1r,3r)-3-(hydroxymethyl)cyclobutoxy)pyridin-1-ium (8 g, 22.8 mmol, 97% yield) was obtained as a white solid.
工程7

エタノール(100mL)中の1-ベンジル-4-((1r,3r)-3-(ヒドロキシメチル)シクロブトキシ)ピリジン-1-イウム(8g、22.84mmol、1当量)の溶液に、水素化ホウ素ナトリウム(1.73g、45.68mmol、2当量)を0℃で添加した。反応混合物を20℃で4時間撹拌した。水(100mL)を混合物に添加して、反応物をクエンチした。得られた混合物を酢酸エチル(100mL×3)で抽出した。合わせた有機相をブライン(100mL)で洗浄し、硫酸ナトリウムで乾燥させ、濾過し、真空中で濃縮した。残渣を、シリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=50:1~0:1)によって精製した。((1r,3r)-3-((1-ベンジル-1,2,3,6-テトラヒドロピリジン-4-イル)オキシ)シクロブチル)メタノール(5.2g、19.02mmol、収率83%)を淡い黄色の油として得た。 To a solution of 1-benzyl-4-((1r,3r)-3-(hydroxymethyl)cyclobutoxy)pyridin-1-ium (8 g, 22.84 mmol, 1 equiv.) in ethanol (100 mL) was added sodium borohydride (1.73 g, 45.68 mmol, 2 equiv.) at 0°C. The reaction mixture was stirred at 20°C for 4 h. Water (100 mL) was added to the mixture to quench the reaction. The resulting mixture was extracted with ethyl acetate (100 mL x 3). The combined organic phase was washed with brine (100 mL), dried over sodium sulfate, filtered, and concentrated in vacuum. The residue was purified by silica gel chromatography (petroleum ether:ethyl acetate = 50:1 to 0:1). ((1r,3r)-3-((1-benzyl-1,2,3,6-tetrahydropyridin-4-yl)oxy)cyclobutyl)methanol (5.2 g, 19.02 mmol, 83% yield) was obtained as a pale yellow oil.
工程8
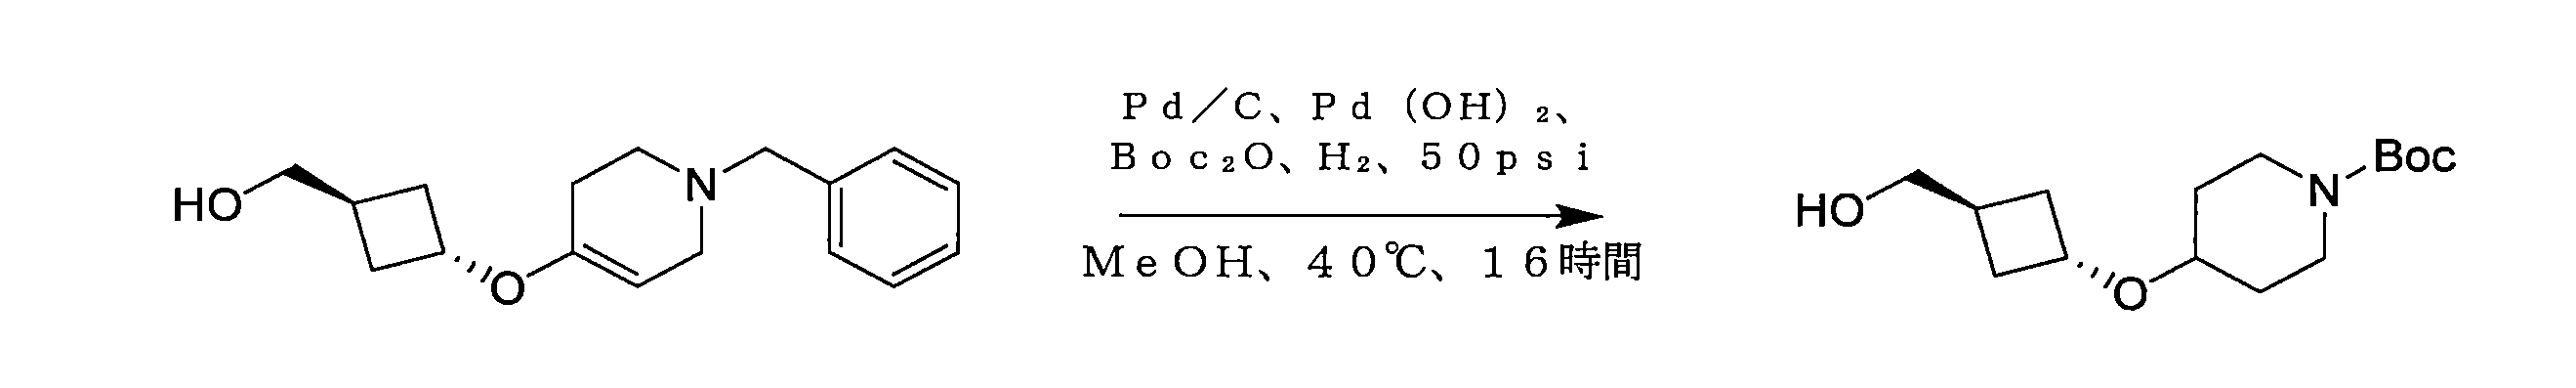
メタノール(100mL)中の[3-[(1-ベンジル-3,6-ジヒドロ-2H-ピリジン-4-イル)オキシ]シクロブチル]メタノール(5.2g、19.02mmol、1当量)およびジ-tert-ブチルジカーボネート(8.30g、38.04mmol、8.7mL、2当量)の溶液に、10%パラジウム活性炭素触媒(0.7g)および水酸化パラジウム20%炭素(0.7g)を窒素下で添加した。反応混合物を水素化(50psi)で、40℃で16時間撹拌した。混合物を、セライトパッドを通して濾過し、濾液を濃縮して、粗生成物を得た。残渣を、シリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=100:1~0:1)によって精製した。tert-ブチル4-((1r,3r)-3-(ヒドロキシメチル)シクロブトキシ)ピペリジン-1-カルボキシレート(4.5g、15.77mmol、収率83%)を無色の油として得た。 To a solution of [3-[(1-benzyl-3,6-dihydro-2H-pyridin-4-yl)oxy]cyclobutyl]methanol (5.2 g, 19.02 mmol, 1 eq.) and di-tert-butyl dicarbonate (8.30 g, 38.04 mmol, 8.7 mL, 2 eq.) in methanol (100 mL), 10% palladium on activated carbon catalyst (0.7 g) and palladium hydroxide 20% on carbon (0.7 g) were added under nitrogen. The reaction mixture was stirred at 40° C. for 16 h under hydrogenation (50 psi). The mixture was filtered through a celite pad and the filtrate was concentrated to give the crude product. The residue was purified by silica gel chromatography (petroleum ether:ethyl acetate=100:1 to 0:1). tert-Butyl 4-((1r,3r)-3-(hydroxymethyl)cyclobutoxy)piperidine-1-carboxylate (4.5 g, 15.77 mmol, 83% yield) was obtained as a colorless oil.
工程9

ジクロロメタン(25mL)中のtert-ブチル4-((1r,3r)-3-(ヒドロキシメチル)シクロブトキシ)ピペリジン-1-カルボキシレート(2.5g、8.76mmol、1当量)の溶液に、デスマーチンペルヨージナン(4.46g、10.51mmol、1.2当量)を0℃で添加した。反応混合物を20℃で2時間撹拌した。混合物を濾過し、飽和重炭酸ナトリウム(50mL)を混合物に添加した。水層を、酢酸エチル(50mL×3)で抽出した。合わせた有機層をブライン(80mL)で洗浄し、硫酸ナトリウムで乾燥させ、濾過し、真空中で濃縮させた。残渣を、シリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=1:0~5:1)によって精製した。tert-ブチル4-((1r,3r)-3-ホルミルシクロブトキシ)ピペリジン-1-カルボキシレート(2.0g、7.06mmol、収率80%)を無色の油として得た。 To a solution of tert-butyl 4-((1r,3r)-3-(hydroxymethyl)cyclobutoxy)piperidine-1-carboxylate (2.5 g, 8.76 mmol, 1 eq.) in dichloromethane (25 mL) was added Dess-Martin periodinane (4.46 g, 10.51 mmol, 1.2 eq.) at 0 °C. The reaction mixture was stirred at 20 °C for 2 h. The mixture was filtered and saturated sodium bicarbonate (50 mL) was added to the mixture. The aqueous layer was extracted with ethyl acetate (50 mL x 3). The combined organic layers were washed with brine (80 mL), dried over sodium sulfate, filtered, and concentrated in vacuo. The residue was purified by silica gel chromatography (petroleum ether:ethyl acetate = 1:0 to 5:1). tert-Butyl 4-((1r,3r)-3-formylcyclobutoxy)piperidine-1-carboxylate (2.0 g, 7.06 mmol, 80% yield) was obtained as a colorless oil.
工程10

メタノール(20mL)中のtert-ブチル4-((1r,3r)-3-ホルミルシクロブトキシ)ピペリジン-1-カルボキシレート(2.0g、7.06mmol、1当量)の溶液に、炭酸カリウム(1.95g、14.12mmol、2当量)および1-ジアゾ-1-ジメトキシホスホリル-プロパン-2-オン(1.42g、7.41mmol、1.05当量)を0℃で添加した。反応混合物を20℃で12時間撹拌した。水(100mL)を混合物に添加し、得られた混合物を酢酸エチル(50mL×3)で抽出した。合わせた有機相をブライン(50mL)で洗浄し、硫酸ナトリウムで乾燥させ、濾過し、真空中で濃縮した。残渣を、シリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=1:0~20:1)によって精製した。tert-ブチル4-((1r,3r)-3-エチニルシクロブトキシ)ピペリジン-1-カルボキシレート(1.3g、4.65mmol、収率66%)を無色の油として得た。 To a solution of tert-butyl 4-((1r,3r)-3-formylcyclobutoxy)piperidine-1-carboxylate (2.0 g, 7.06 mmol, 1 equiv.) in methanol (20 mL) was added potassium carbonate (1.95 g, 14.12 mmol, 2 equiv.) and 1-diazo-1-dimethoxyphosphoryl-propan-2-one (1.42 g, 7.41 mmol, 1.05 equiv.) at 0 °C. The reaction mixture was stirred at 20 °C for 12 h. Water (100 mL) was added to the mixture, and the resulting mixture was extracted with ethyl acetate (50 mL × 3). The combined organic phase was washed with brine (50 mL), dried over sodium sulfate, filtered, and concentrated in vacuum. The residue was purified by silica gel chromatography (petroleum ether: ethyl acetate = 1:0 to 20:1). tert-Butyl 4-((1r,3r)-3-ethynylcyclobutoxy)piperidine-1-carboxylate (1.3 g, 4.65 mmol, 66% yield) was obtained as a colorless oil.
工程11
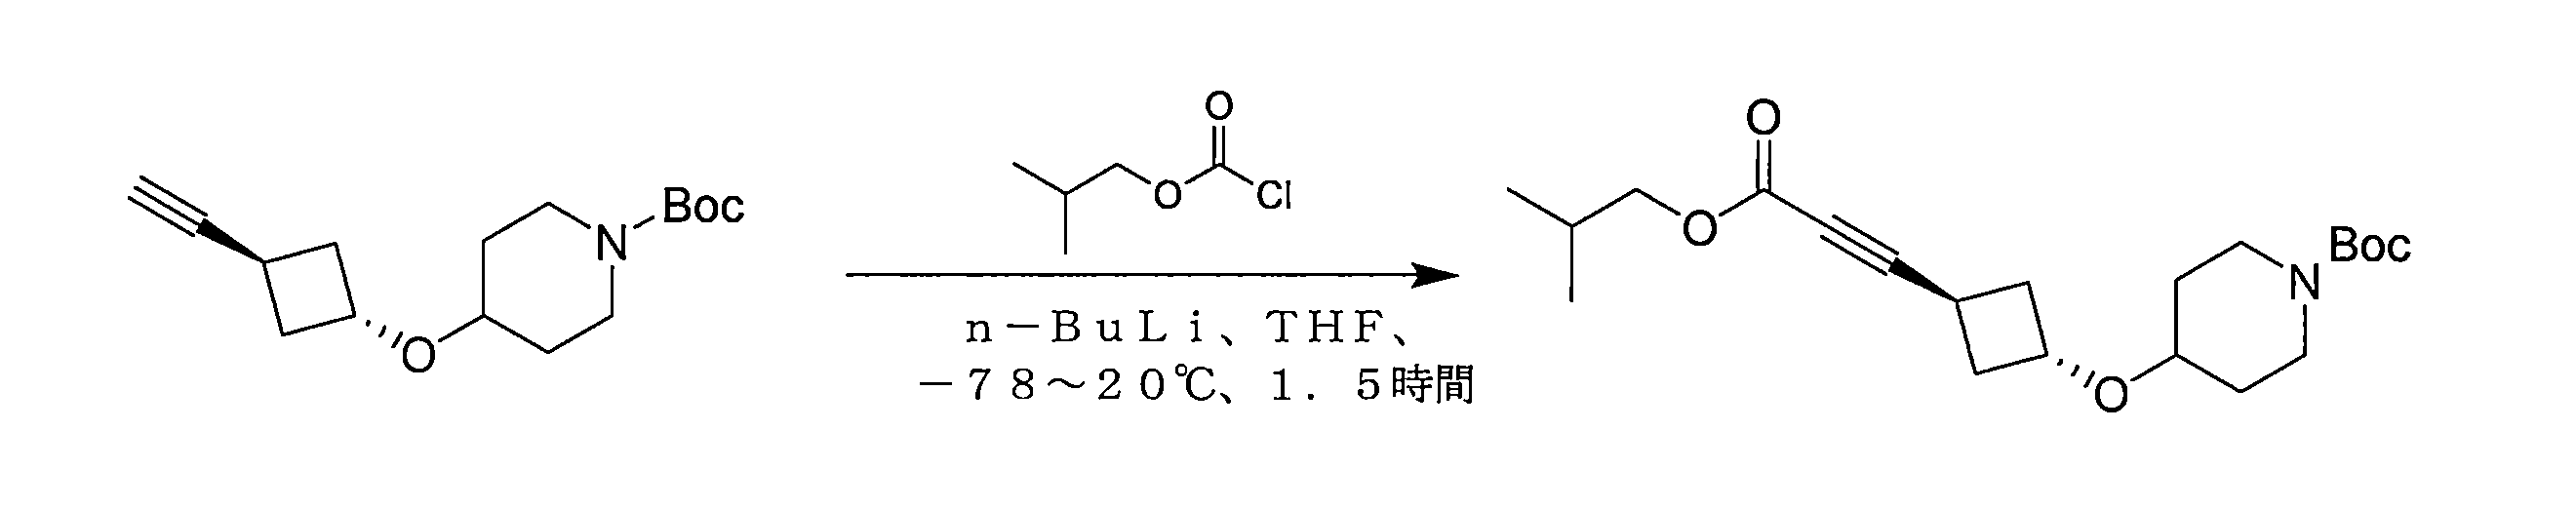
テトラヒドロフラン(17mL)中のtert-ブチル4-((1r,3r)-3-エチニルシクロブトキシ)ピペリジン-1-カルボキシレート(1.7g、6.09mmol、1当量)の溶液に、n-ブチルリチウム(2.5M、3.6mL、1.5当量)を-78℃で添加した。混合物を-78℃で0.5時間撹拌した。次いで、クロロギ酸イソブチル(1.66g、12.17mmol、1.6mL、2当量)を、混合物に添加し、反応混合物を-78℃で0.5時間、および20℃で1時間撹拌した。飽和塩化アンモニウム(80mL)を混合物に添加し、得られた混合物を酢酸エチル(80mL×3)で抽出した。合わせた有機相をブライン(100mL)で洗浄し、硫酸ナトリウムで乾燥させ、濾過し、真空中で濃縮した。残渣を、シリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=1:0~5:1)によって精製した。tert-ブチル4-((1r,3r)-3-(3-イソブトキシ-3-オキソプロプ-1-イン-1-イル)シクロブトキシ)ピペリジン-1-カルボキシレート(2g、5.27mmol、収率86%)を淡い黄色の油として得た。 To a solution of tert-butyl 4-((1r,3r)-3-ethynylcyclobutoxy)piperidine-1-carboxylate (1.7 g, 6.09 mmol, 1 eq.) in tetrahydrofuran (17 mL) was added n-butyllithium (2.5 M, 3.6 mL, 1.5 eq.) at −78°C. The mixture was stirred at −78°C for 0.5 h. Then, isobutyl chloroformate (1.66 g, 12.17 mmol, 1.6 mL, 2 eq.) was added to the mixture, and the reaction mixture was stirred at −78°C for 0.5 h and at 20°C for 1 h. Saturated ammonium chloride (80 mL) was added to the mixture, and the resulting mixture was extracted with ethyl acetate (80 mL x 3). The combined organic phase was washed with brine (100 mL), dried over sodium sulfate, filtered, and concentrated in vacuum. The residue was purified by silica gel chromatography (petroleum ether:ethyl acetate = 1:0 to 5:1). tert-Butyl 4-((1r,3r)-3-(3-isobutoxy-3-oxoprop-1-yn-1-yl)cyclobutoxy)piperidine-1-carboxylate (2 g, 5.27 mmol, 86% yield) was obtained as a pale yellow oil.
工程12

ジクロロメタン(20mL)中のtert-ブチル4-[3-(3-イソブトキシ-3-オキソ-プロパ-1-イニル)シクロブトキシ]ピペリジン-1-カルボキシレート(2g、5.27mmol、1当量)の溶液に、DCM(1M、6.3mL、1.2当量)中の水素化ジイソブチルアルミニウム溶液を-78℃で添加した。反応混合物を-78℃で1時間撹拌した。飽和重炭酸ナトリウム(10mL)を混合物に添加し、酢酸エチル(100mL)を混合物に添加した。混合物をセライトパッドを通して濾過し、酢酸エチル(50mL×3)で抽出した。合わせた有機相をブライン(70mL)で洗浄し、硫酸ナトリウムで乾燥させ、濾過し、真空中で濃縮した。残渣を、シリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=1:0~5:1)によって精製した。tert-ブチル4-((1r,3r)-3-(3-オキソプロプ-1-イン-1-イル)シクロブトキシ)ピペリジン-1-カルボキシレート(1.2g、3.90mmol、収率74%)を無色の油として得た。 To a solution of tert-butyl 4-[3-(3-isobutoxy-3-oxo-prop-1-ynyl)cyclobutoxy]piperidine-1-carboxylate (2 g, 5.27 mmol, 1 eq.) in dichloromethane (20 mL) was added a solution of diisobutylaluminum hydride in DCM (1 M, 6.3 mL, 1.2 eq.) at −78 °C. The reaction mixture was stirred at −78 °C for 1 h. Saturated sodium bicarbonate (10 mL) was added to the mixture, and ethyl acetate (100 mL) was added to the mixture. The mixture was filtered through a celite pad and extracted with ethyl acetate (50 mL × 3). The combined organic phase was washed with brine (70 mL), dried over sodium sulfate, filtered, and concentrated in vacuum. The residue was purified by silica gel chromatography (petroleum ether:ethyl acetate = 1:0 to 5:1). tert-Butyl 4-((1r,3r)-3-(3-oxoprop-1-yn-1-yl)cyclobutoxy)piperidine-1-carboxylate (1.2 g, 3.90 mmol, 74% yield) was obtained as a colorless oil.
工程13

エタノール(10mL)中のtert-ブチル4-[3-(3-オキソプロパ-1-イニル)シクロブトキシ]ピペリジン-1-カルボキシレート(1.2g、3.90mmol、1当量)の溶液に、水素化ホウ素ナトリウム(177mg、4.68mmol、1.2当量)を0℃で、3回に分けて添加した。反応混合物を20℃で1時間撹拌した。水(100mL)を混合物に加え、得られた混合物を酢酸エチル(50mL×3)で抽出した。合わせた有機相をブライン(50mL)で洗浄し、硫酸ナトリウムで乾燥させ、濾過し、真空中で濃縮した。残渣を、シリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=50:1~1:1)によって精製した。tert-ブチル4-((1r,3r)-3-(3-ヒドロキシプロプ-1-イン-1-イル)シクロブトキシ)ピペリジン-1-カルボキシレート(1.06g、3.43mmol、収率87%)を無色の油として得た。 To a solution of tert-butyl 4-[3-(3-oxoprop-1-ynyl)cyclobutoxy]piperidine-1-carboxylate (1.2 g, 3.90 mmol, 1 equiv.) in ethanol (10 mL) was added sodium borohydride (177 mg, 4.68 mmol, 1.2 equiv.) in three portions at 0 °C. The reaction mixture was stirred at 20 °C for 1 h. Water (100 mL) was added to the mixture, and the resulting mixture was extracted with ethyl acetate (50 mL x 3). The combined organic phase was washed with brine (50 mL), dried over sodium sulfate, filtered, and concentrated in vacuum. The residue was purified by silica gel chromatography (petroleum ether:ethyl acetate = 50:1 to 1:1). tert-Butyl 4-((1r,3r)-3-(3-hydroxyprop-1-yn-1-yl)cyclobutoxy)piperidine-1-carboxylate (1.06 g, 3.43 mmol, 87% yield) was obtained as a colorless oil.
工程14

ジクロロメタン(10mL)中のtert-ブチル4-((1r,3r)-3-(3-ヒドロキシプロパ-1-イン-1-イル)シクロブトキシ)ピペリジン-1-カルボキシレート(1.06g、3.43mmol、1当量)の溶液に、トリフェニルホスフィン(1.08g、4.11mmol、1.2当量)および四臭化炭素(1.36g、4.11mmol、1.2当量)を0℃で添加した。反応混合物を20℃で3時間撹拌した。混合物を真空中で濃縮した。残渣を、シリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=1:0~20:1)によって精製した。tert-ブチル4-[3-(3-ブロモプロプ-1-イニル)シクロブトキシ)ピペリジン-1-カルボキシレート(1g、2.69mmol、収率78%)を淡い黄色の油として得た。 To a solution of tert-butyl 4-((1r,3r)-3-(3-hydroxyprop-1-yn-1-yl)cyclobutoxy)piperidine-1-carboxylate (1.06 g, 3.43 mmol, 1 equiv.) in dichloromethane (10 mL) was added triphenylphosphine (1.08 g, 4.11 mmol, 1.2 equiv.) and carbon tetrabromide (1.36 g, 4.11 mmol, 1.2 equiv.) at 0 °C. The reaction mixture was stirred at 20 °C for 3 h. The mixture was concentrated in vacuum. The residue was purified by silica gel chromatography (petroleum ether: ethyl acetate = 1:0 to 20:1). tert-butyl 4-[3-(3-bromoprop-1-ynyl)cyclobutoxy)piperidine-1-carboxylate (1 g, 2.69 mmol, 78% yield) was obtained as a pale yellow oil.
上記の他の実施例について記載される手順を使用して、以下のスキームに従って、tert-ブチル4-((1r,3r)-3-(3-ヒドロキシプロパ-1-イン-1-イル)シクロブトキシ)ピペリジン-1-カルボキシレートを最終化合物に変換した。
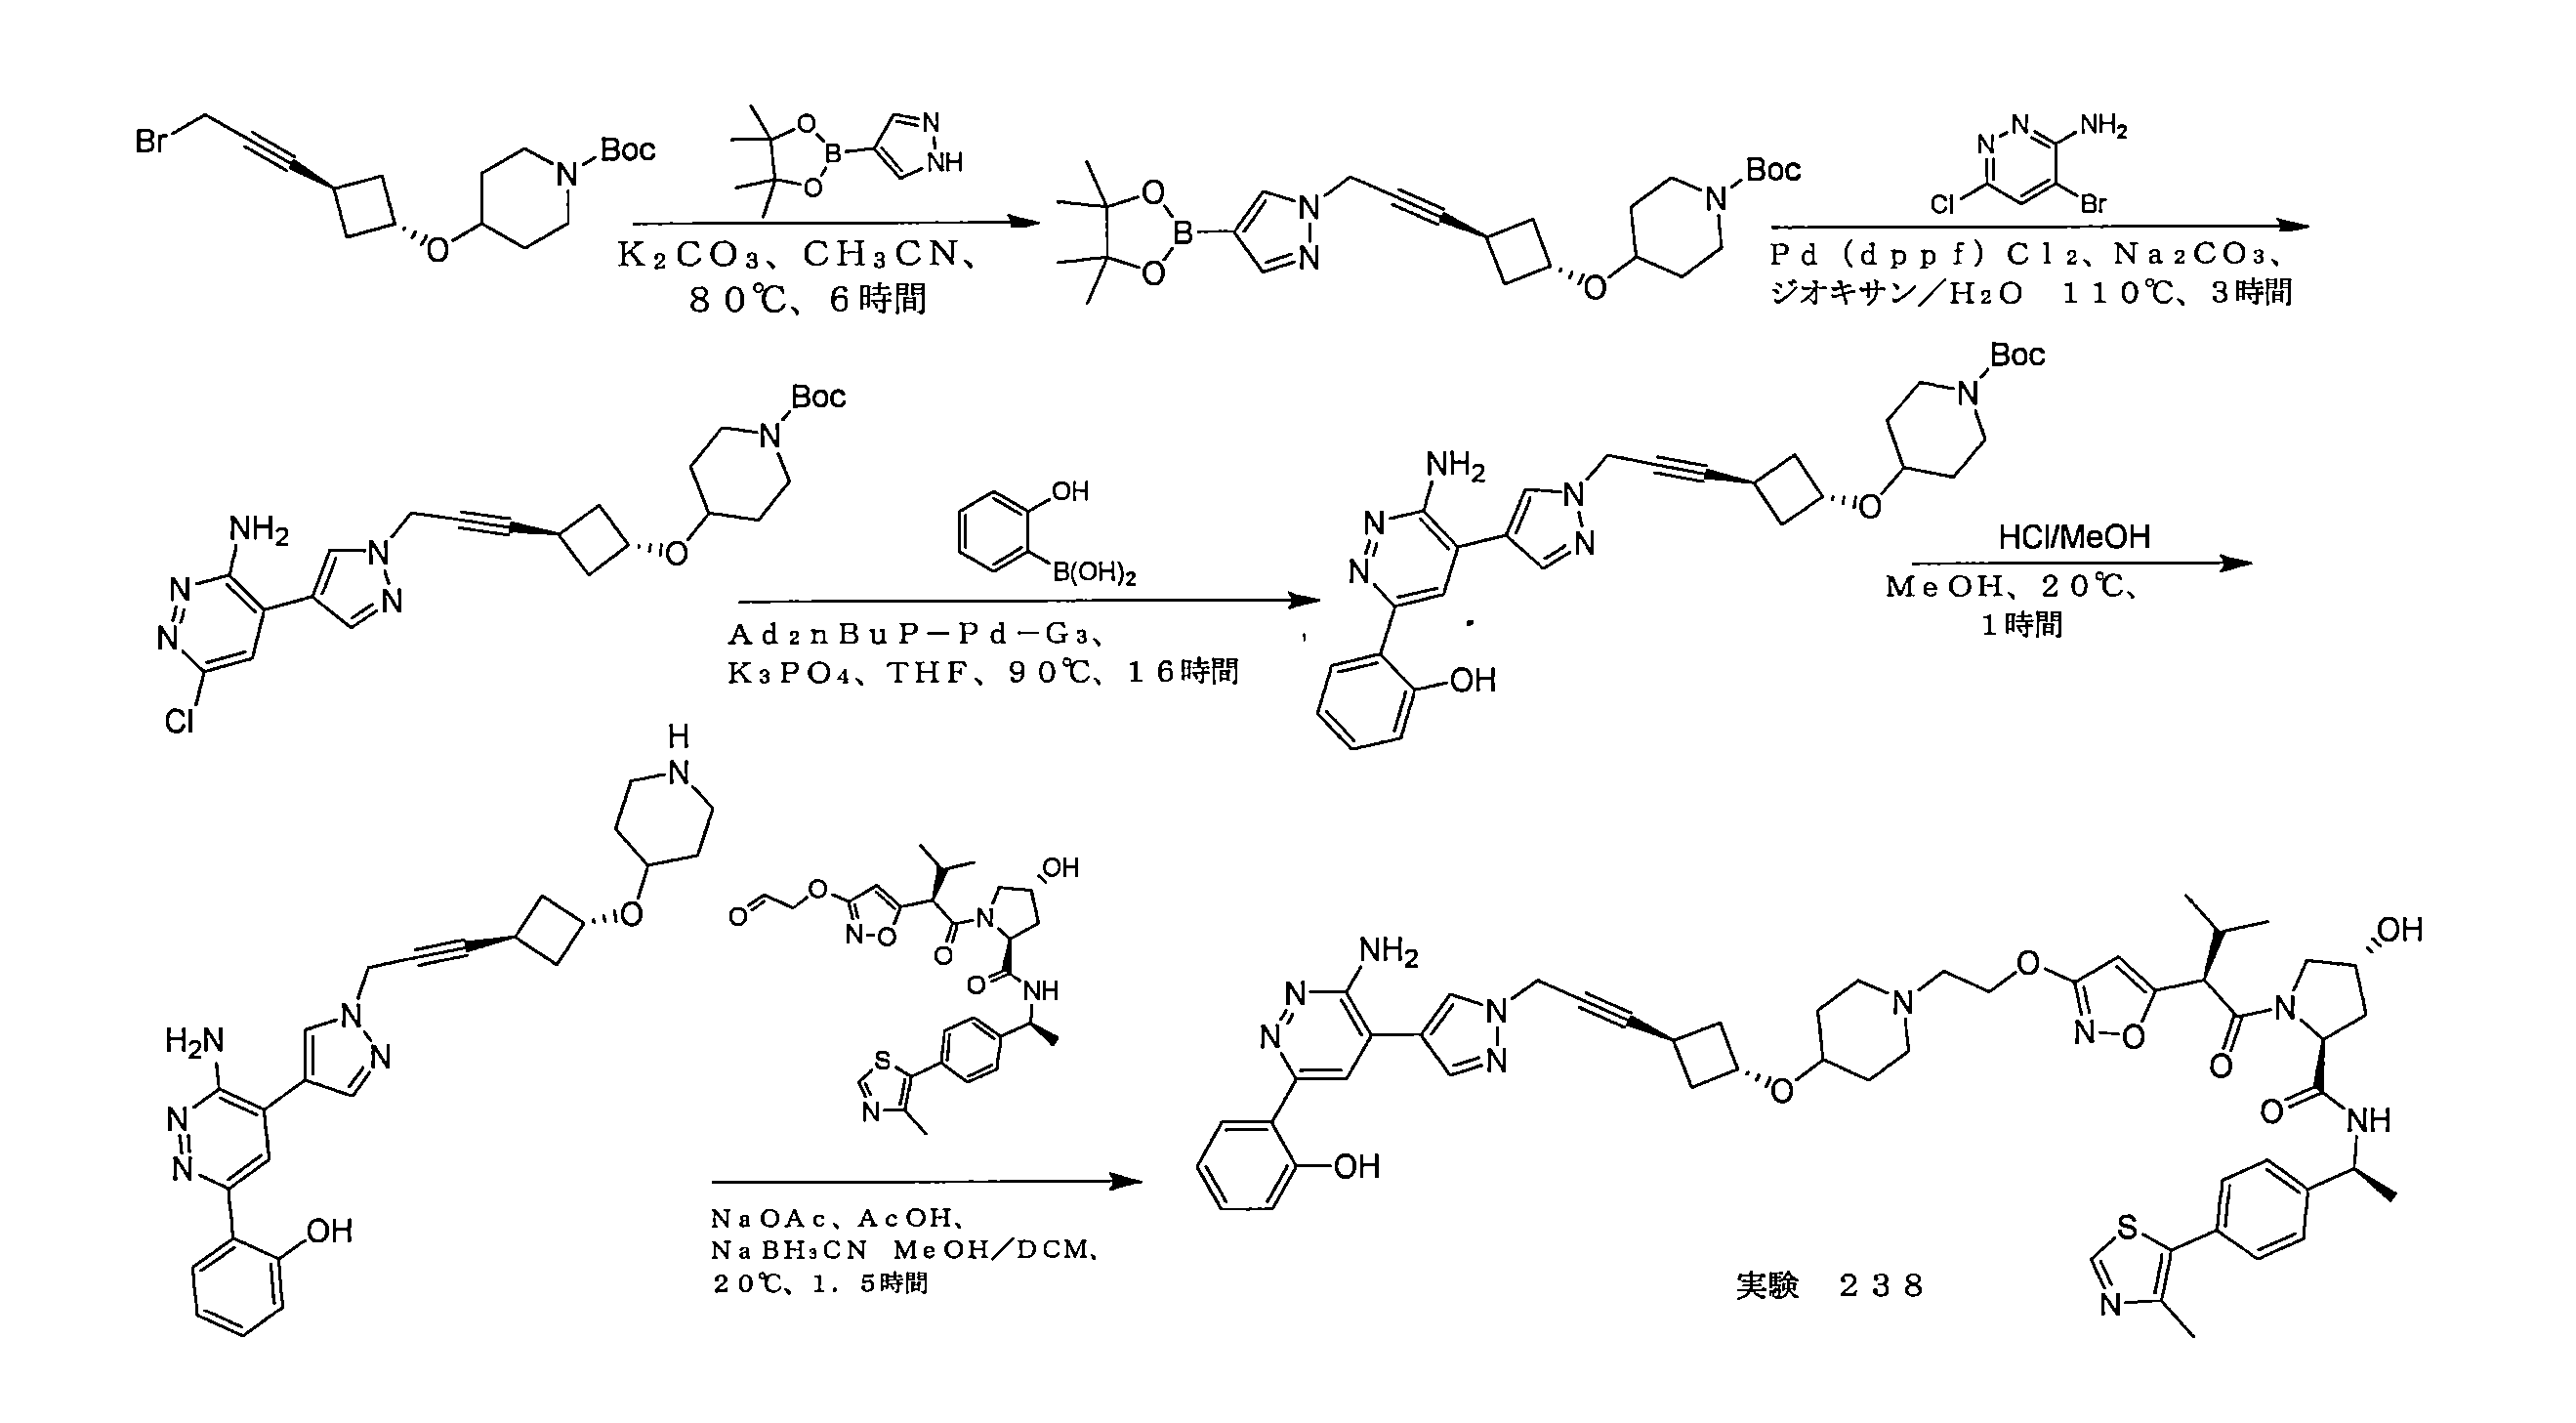
例示的な化合物216の例示的な合成
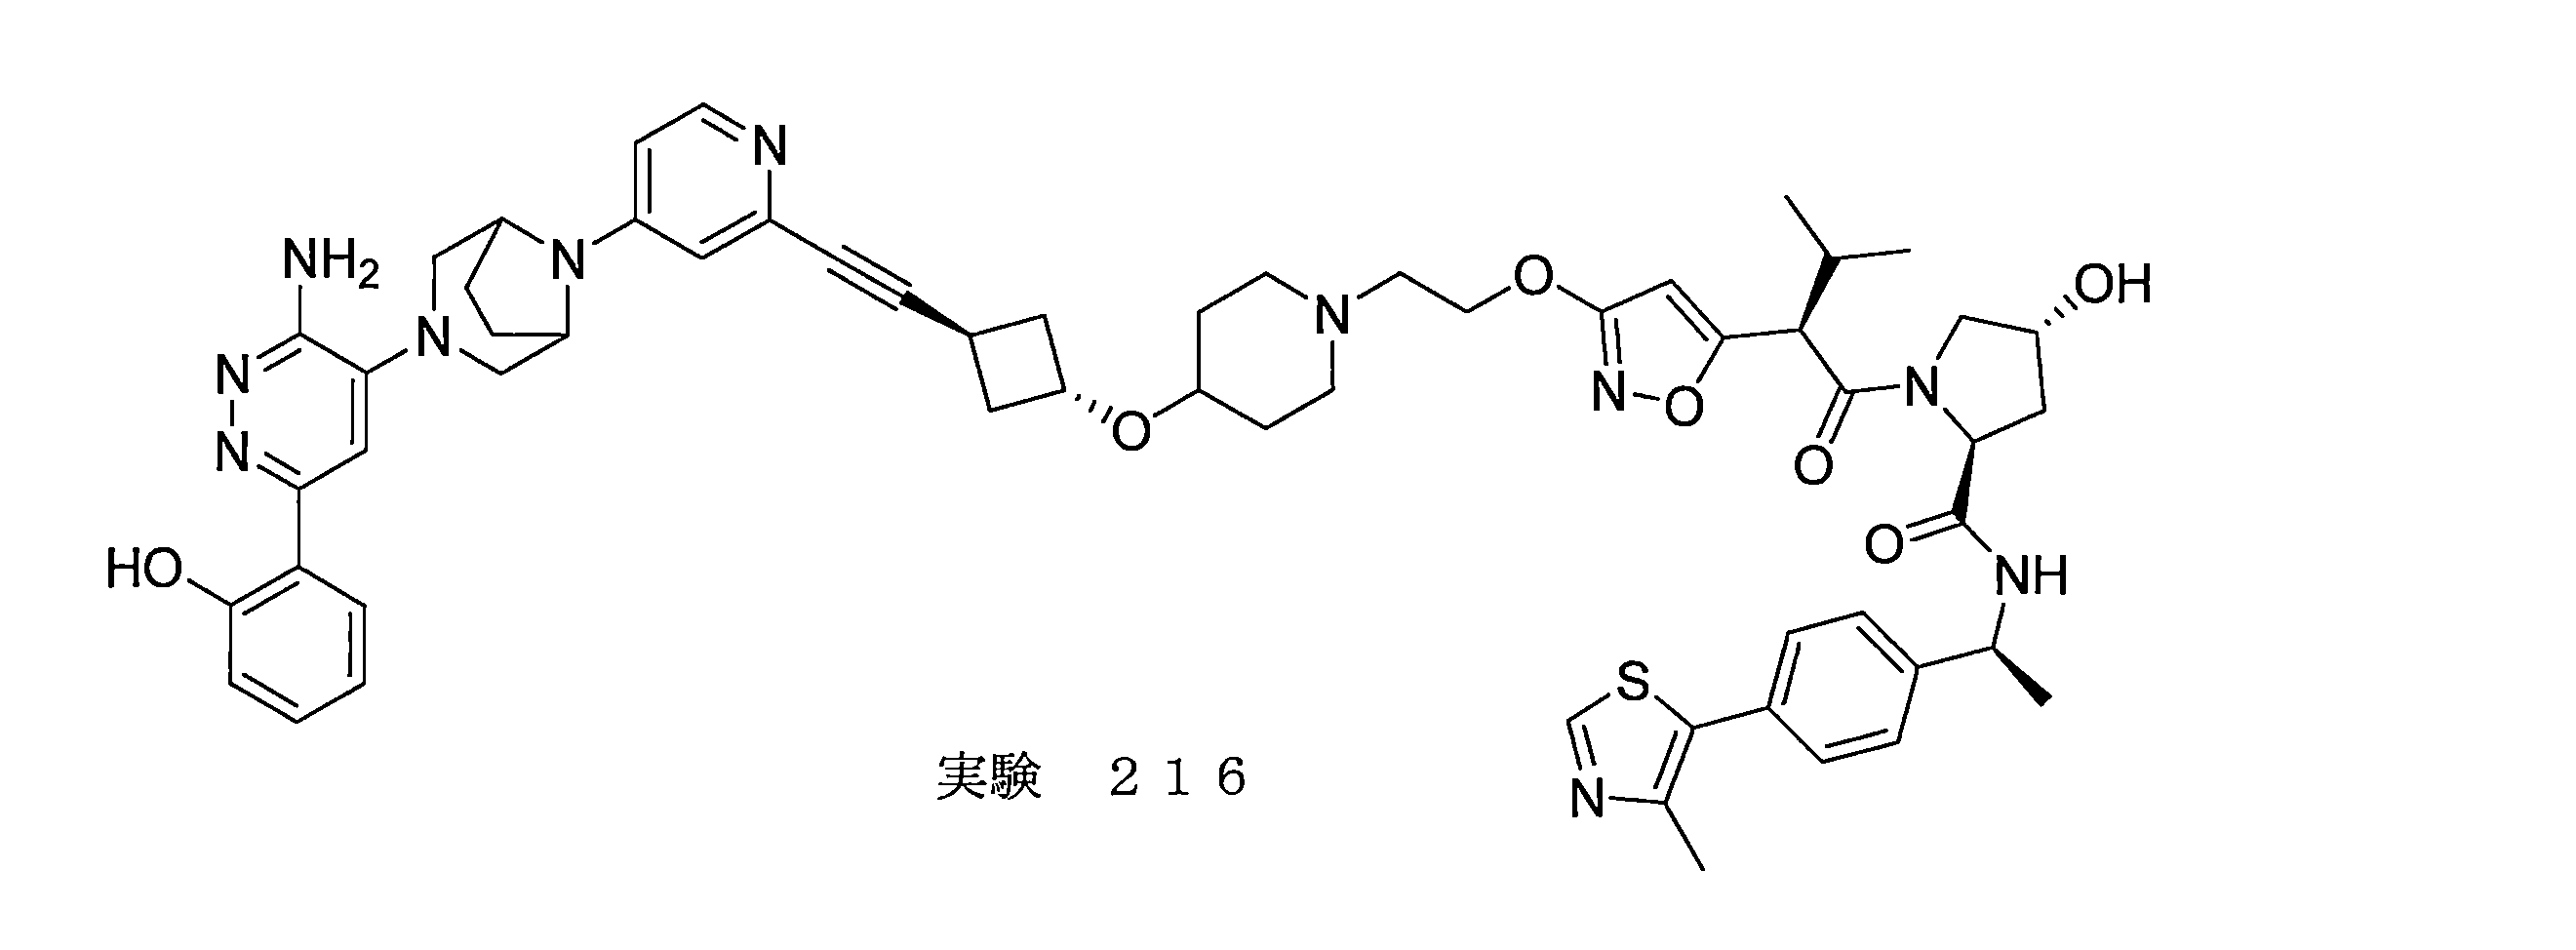
工程1

ジメチルスルホキシド(30mL)中の2-ブロモ-4-フルオロ-ピリジン(310mg、1.76mmol、1.2当量)、4-(3,8-ジアザビシクロ[3.2.1]オクタン-3-イル)-6-[2-(メトキシメトキシ)フェニル]ピリダジン-3-アミン(500mg、1.46mmol、1当量)、およびジイソプロピルエチルアミン(2.27g、17.57mmol、3mL、12当量)の混合物を脱気し、窒素で3回パージし、次いで、混合物を窒素雰囲気下で、130℃で5時間撹拌した。反応混合物を水(50mL)で希釈し、酢酸エチル(50mL)で3回抽出した。合わせた有機層を無水硫酸ナトリウムで乾燥させ、濃縮して、粗生成物を得た。残渣を、半分取逆相クロマトグラフィー(カラム:Phenomenex Synergi C18 150*30mm*4um;移動相:[水(0.225%FA)-ACN];B%:15%~45%、10分)によって精製した。4-[8-(2-ブロモ-4-ピリジル)-3,8-ジアザビシクロ[3.2.1]オクタン-3-イル]-6-[2-(メトキシメトキシ)フェニル]ピリダジン-3-アミン(562mg、1.13mmol、収率77%)を黄色の固体として得た。 A mixture of 2-bromo-4-fluoro-pyridine (310 mg, 1.76 mmol, 1.2 eq.), 4-(3,8-diazabicyclo[3.2.1]octan-3-yl)-6-[2-(methoxymethoxy)phenyl]pyridazin-3-amine (500 mg, 1.46 mmol, 1 eq.), and diisopropylethylamine (2.27 g, 17.57 mmol, 3 mL, 12 eq.) in dimethylsulfoxide (30 mL) was degassed and purged with nitrogen three times, then the mixture was stirred under nitrogen atmosphere at 130 °C for 5 h. The reaction mixture was diluted with water (50 mL) and extracted three times with ethyl acetate (50 mL). The combined organic layers were dried over anhydrous sodium sulfate and concentrated to give the crude product. The residue was purified by semi-preparative reverse phase chromatography (column: Phenomenex Synergi C18 150*30mm*4um; mobile phase: [water (0.225% FA)-ACN]; B%: 15%-45%, 10 min). 4-[8-(2-bromo-4-pyridyl)-3,8-diazabicyclo[3.2.1]octan-3-yl]-6-[2-(methoxymethoxy)phenyl]pyridazin-3-amine (562 mg, 1.13 mmol, 77% yield) was obtained as a yellow solid.
工程2

ジメチルホルムアミド(35mL)中の4-[8-(2-ブロモ-4-ピリジル)-3,8-ジアザビシクロ[3.2.1]オクタン-3-イル]-6-[2-(メトキシメトキシ)フェニル]ピリダジン-3-アミン(568mg、1.14mmol、1当量)、tert-ブチル4-((1r,3r)-3-エチニルシクロブトキシ)ピペリジン-1-カルボキシレート(479mg、1.71mmol、1.5当量)、ビス(トリフェニルホスフィン)パラジウム(II)二塩化物(80mg、0.1mmol、0.1当量)、ヨウ化銅(22mg、0.1mmol、0.1当量)、およびN-イソプロピルプロパン-2-アミン(8.13g、80.38mmol、11.36mL、70.39当量)の混合物を脱気し、窒素で3回パージし、次いで、混合物を窒素雰囲気下で、75℃で12時間撹拌した。反応混合物を水(30mL)で希釈し、酢酸エチル(60mL)で3回抽出した。合わせた有機層をブライン(50mL)で2回洗浄し、無水硫酸ナトリウムで乾燥させ、濃縮して、粗生成物を得た。残渣を、半分取逆相クロマトグラフィー(カラム:Phenomenex luna C18 250*50mm*10um;移動相:[水(0.225%FA)-ACN];B%:15%~45%、25分)によって精製した。tert-ブチル4-((1r,3r)-3-((4-(3-(3-アミノ-6-(2-(メトキシメトキシ)フェニル)ピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル)エチニル)シクロブトキシ)ピペリジン-1-カルボキシレート(581mg、0.8mmol、収率73%)を黄色の固体として得た。 4-[8-(2-bromo-4-pyridyl)-3,8-diazabicyclo[3.2.1]octan-3-yl]-6-[2-(methoxymethoxy)phenyl]pyridazin-3-amine (568 mg, 1.14 mmol, 1 equiv.), tert-butyl 4-((1r,3r)-3-ethynylcyclobutoxy)piperidine-1-carboxylate (479 mg, 1.71 mmol) in dimethylformamide (35 mL) A mixture of 1,2-dichlorophenyl ether (1.5 equiv.), bis(triphenylphosphine)palladium(II) dichloride (80 mg, 0.1 mmol, 0.1 equiv.), copper iodide (22 mg, 0.1 mmol, 0.1 equiv.), and N-isopropylpropan-2-amine (8.13 g, 80.38 mmol, 11.36 mL, 70.39 equiv.) was degassed and purged with nitrogen three times, then the mixture was stirred under nitrogen atmosphere at 75° C. for 12 h. The reaction mixture was diluted with water (30 mL) and extracted three times with ethyl acetate (60 mL). The combined organic layers were washed twice with brine (50 mL), dried over anhydrous sodium sulfate, and concentrated to give the crude product. The residue was purified by semi-preparative reverse phase chromatography (column: Phenomenex luna C18 250*50mm*10um; mobile phase: [water (0.225% FA)-ACN]; B%: 15%-45%, 25 min). tert-Butyl 4-((1r,3r)-3-((4-(3-(3-amino-6-(2-(methoxymethoxy)phenyl)pyridazin-4-yl)-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl)ethynyl)cyclobutoxy)piperidine-1-carboxylate (581 mg, 0.8 mmol, 73% yield) was obtained as a yellow solid.
上記の他の実施例について記載されるように、tert-ブチル4-((1r,3r)-3-((4-(3-(3-アミノ-6-(2-(メトキシメトキシ)フェニル)ピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル)エチニル)シクロブトキシ)ピペリジン-1-カルボキシレートを最終化合物に変換した。 tert-Butyl 4-((1r,3r)-3-((4-(3-(3-amino-6-(2-(methoxymethoxy)phenyl)pyridazin-4-yl)-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl)ethynyl)cyclobutoxy)piperidine-1-carboxylate was converted to the final compound as described for other examples above.
例示的な化合物217および218の例示的な合成
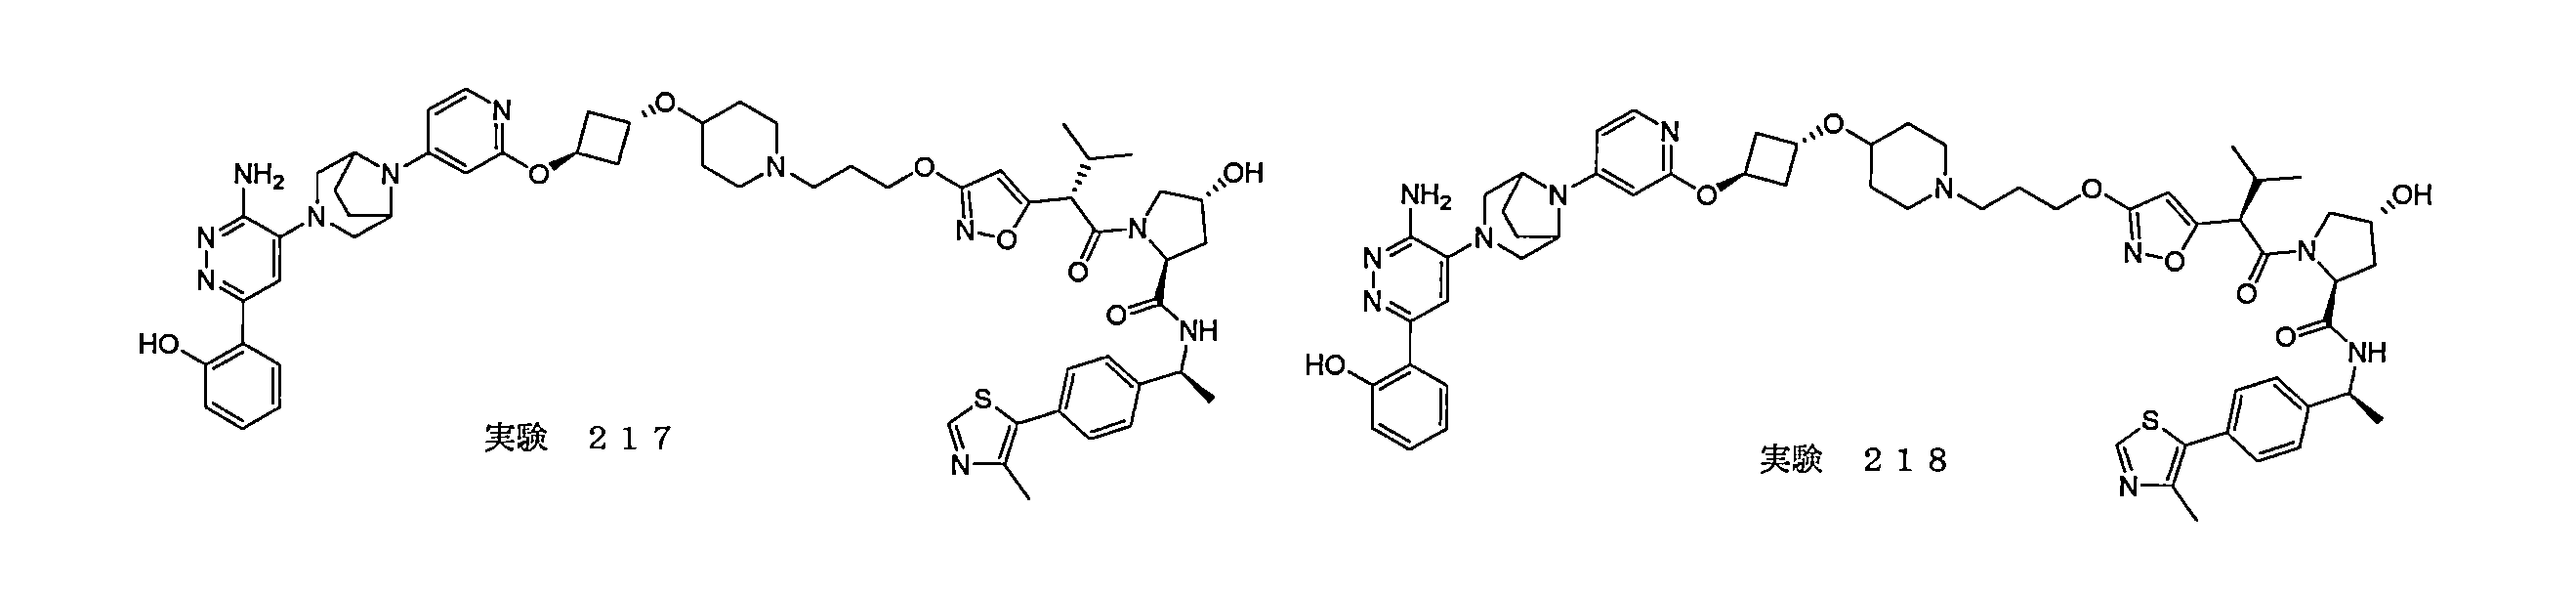
工程1
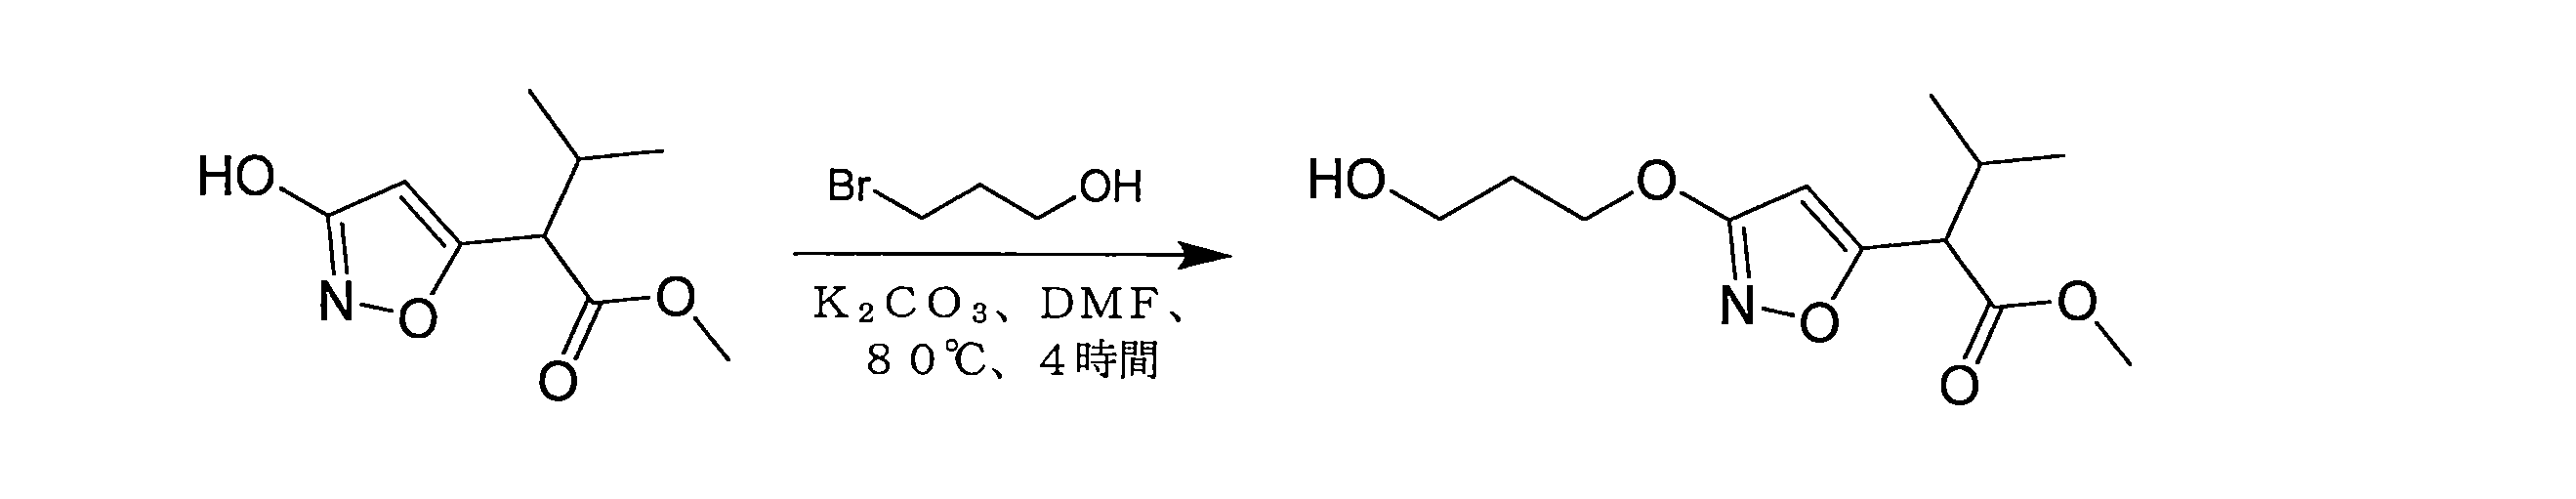
N,N-ジメチルホルムアミド(5mL)中のメチル2-(3-ヒドロキシイソキサゾール-5-イル)-3-メチル-ブタノエート(500mg、2.51mmol、1当量)および3-ブロモプロパン-1-オール(523mg、3.77mmol、0.34mL、1.5当量)の溶液に、炭酸カリウム(693mg、5.02mmol、2当量)を20℃で添加し、混合物を80℃で4時間撹拌した。反応混合物を水(20mL)によってクエンチし、次いで、酢酸エチル(10mL×3)で抽出した。合わせた有機層をブライン(10mL)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、減圧下で濃縮して、残渣を得た。残渣を、シリカゲルカラムクロマトグラフィー(石油エーテル/酢酸エチル=20:1~4:1)によって精製した。メチル2-[3-(3-ヒドロキシプロパオキシ)イソキサゾール-5-イル]-3-メチル-ブタノエート(370mg、1.44mmol、収率57%)を無色の油として得た。 To a solution of methyl 2-(3-hydroxyisoxazol-5-yl)-3-methyl-butanoate (500 mg, 2.51 mmol, 1 eq.) and 3-bromopropan-1-ol (523 mg, 3.77 mmol, 0.34 mL, 1.5 eq.) in N,N-dimethylformamide (5 mL), potassium carbonate (693 mg, 5.02 mmol, 2 eq.) was added at 20°C, and the mixture was stirred at 80°C for 4 h. The reaction mixture was quenched by water (20 mL) and then extracted with ethyl acetate (10 mL x 3). The combined organic layers were washed with brine (10 mL), dried over anhydrous sodium sulfate, filtered, and concentrated under reduced pressure to give a residue. The residue was purified by silica gel column chromatography (petroleum ether/ethyl acetate = 20:1 to 4:1). Methyl 2-[3-(3-hydroxypropoxy)isoxazol-5-yl]-3-methyl-butanoate (370 mg, 1.44 mmol, 57% yield) was obtained as a colorless oil.
工程2

ジクロロメタン(5mL)中のメチル2-[3-(3-ヒドロキシプロポキシ)イソキサゾール-5-イル]-3-メチル-ブタノエート(370mg、1.44mmol、1当量)の溶液に、トリエチルアミン(291mg、2.88mmol、0.40mL、2当量)およびp-トルエンスルホニルクロリド(411mg、2.16mmol、1.5当量)を、20℃で逐次的に添加し、20℃で16時間撹拌した。反応混合物を濃縮して、残渣を得た。残渣を、分取TLC(石油エーテル/酢酸エチル=3/1)によって精製した。メチル3-メチル-2-[3-[3-(p-トリルスルホニルオキシ)プロポキシ]イソキサゾール-5-イル]ブタノエート(438mg、1.06mmol、収率74%)を無色の油として得た。 To a solution of methyl 2-[3-(3-hydroxypropoxy)isoxazol-5-yl]-3-methyl-butanoate (370 mg, 1.44 mmol, 1 equiv.) in dichloromethane (5 mL), triethylamine (291 mg, 2.88 mmol, 0.40 mL, 2 equiv.) and p-toluenesulfonyl chloride (411 mg, 2.16 mmol, 1.5 equiv.) were added successively at 20°C and stirred at 20°C for 16 h. The reaction mixture was concentrated to give a residue. The residue was purified by preparative TLC (petroleum ether/ethyl acetate=3/1). Methyl 3-methyl-2-[3-[3-(p-tolylsulfonyloxy)propoxy]isoxazol-5-yl]butanoate (438 mg, 1.06 mmol, 74% yield) was obtained as a colorless oil.
工程3

ジメチルスルホキシド(3mL)中のメチル3-メチル-2-[3-[3-(p-トリルスルホニルオキシ)プロポキシ]イソキサゾール-5-イル]ブタノエート(250mg、0.61mmol、1当量)および2-[6-アミノ-5-[8-[2-[3-(4-ピペリジルオキシ)シクロブトキシ]-4-ピリジル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-イル]ピリダジン-3-イル]フェノール(386mg、0.61mmol、1当量、2トリフルオロ酢酸塩)の溶液に、N,N-ジイソプロピルエチルアミン(314mg、2.43mmol、0.42mL、4当量)およびヨウ化カリウム(10mg、0.06mmol、0.1当量)を20℃で添加し、混合物を80℃で12時間撹拌した。反応混合物を水(20mL)で希釈し、次いで、酢酸エチル(10mL×3)で抽出した。合わせた有機層をブライン(10mL)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、減圧下で濃縮して、残渣を得た。残渣を、シリカゲルカラムクロマトグラフィー(ジクロロメタン/メタノール=10/1)によって精製した。メチル2-[3-[3-[4-[3-[[4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]-2-ピリジル]オキシ]シクロブトキシ]-1-ピペリジル]プロポキシ]イソキサゾール-5-イル]-3-メチル-ブタノエート(90mg、0.11mmol、収率17%)を黄色の油として得た。 To a solution of methyl 3-methyl-2-[3-[3-(p-tolylsulfonyloxy)propoxy]isoxazol-5-yl]butanoate (250 mg, 0.61 mmol, 1 eq.) and 2-[6-amino-5-[8-[2-[3-(4-piperidyloxy)cyclobutoxy]-4-pyridyl]-3,8-diazabicyclo[3.2.1]octan-3-yl]pyridazin-3-yl]phenol (386 mg, 0.61 mmol, 1 eq., 2-trifluoroacetate salt) in dimethyl sulfoxide (3 mL), N,N-diisopropylethylamine (314 mg, 2.43 mmol, 0.42 mL, 4 eq.) and potassium iodide (10 mg, 0.06 mmol, 0.1 eq.) were added at 20°C and the mixture was stirred at 80°C for 12 h. The reaction mixture was diluted with water (20 mL) and then extracted with ethyl acetate (10 mL x 3). The combined organic layers were washed with brine (10 mL), dried over anhydrous sodium sulfate, filtered, and concentrated under reduced pressure to give a residue. The residue was purified by silica gel column chromatography (dichloromethane/methanol = 10/1). Methyl 2-[3-[3-[4-[3-[[4-[3-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl]-2-pyridyl]oxy]cyclobutoxy]-1-piperidyl]propoxy]isoxazol-5-yl]-3-methyl-butanoate (90 mg, 0.11 mmol, 17% yield) was obtained as a yellow oil.
上記の他の実施例について記載される手順を使用して、以下のスキームに従って、メチル2-[3-[3-[4-[3-[[4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]-2-ピリジル]オキシ]シクロブトキシ]-1-ピペリジル]プロポキシ]イソキサゾール-5-イル]-3-メチル-ブタノエートを最終化合物に変換した。
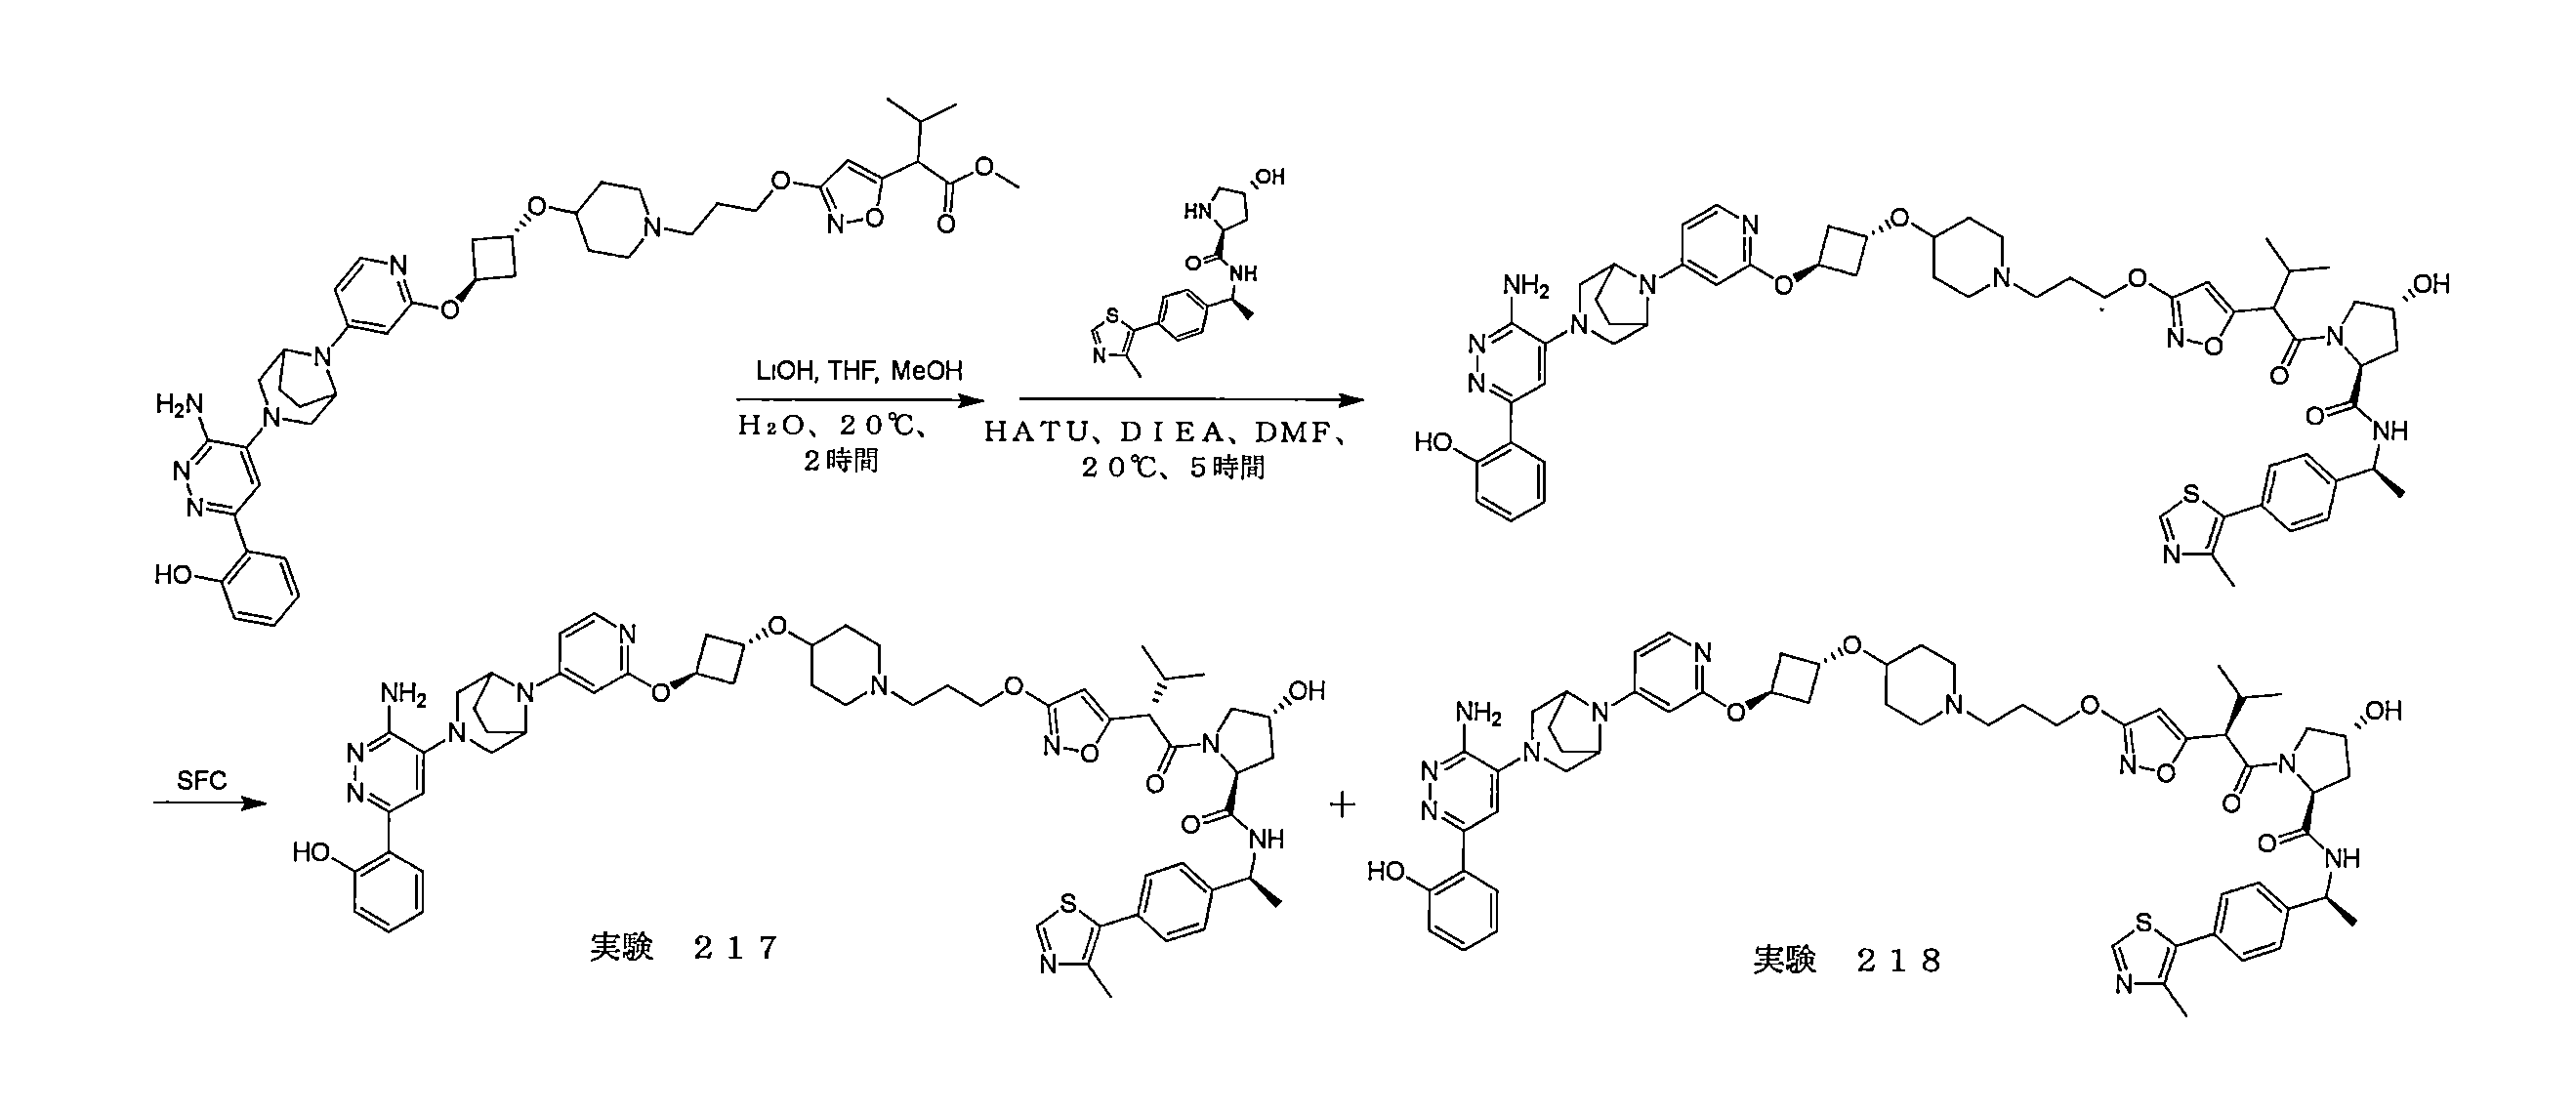
例示的な化合物223の例示的な合成
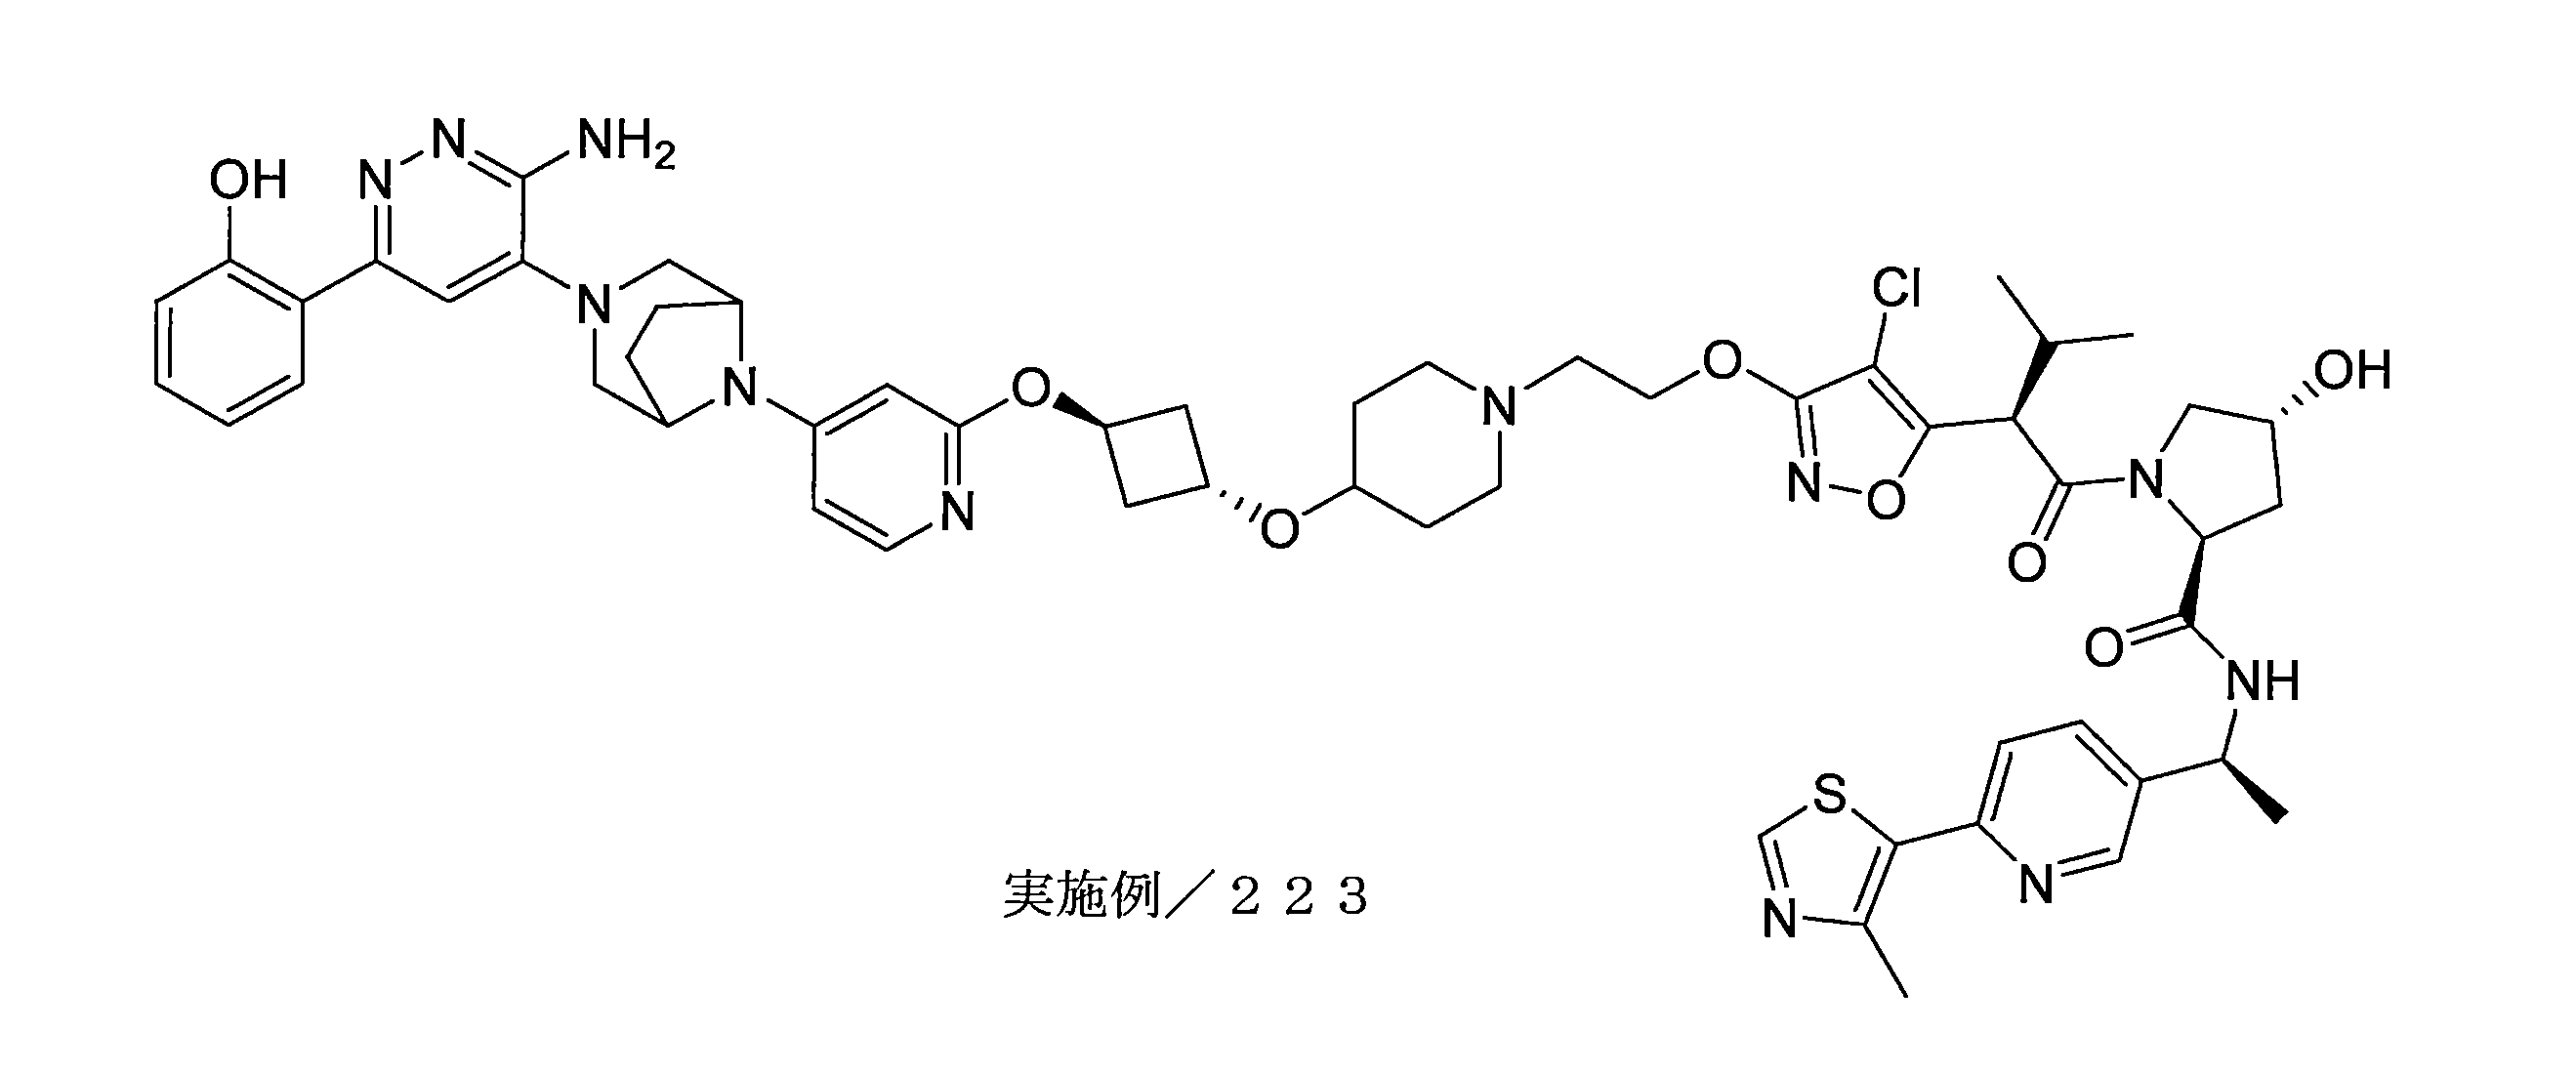
工程1

N,N-ジメチルホルムアミド(10mL)中のメチル2-[3-(2,2-ジエトキシエトキシ)イソキサゾール-5-イル]-3-メチル-ブタノエート(500mg、1.59mmol、1当量)の溶液に、N-クロロスクシンイミド(423mg、3.17mmol、2当量)を添加した。反応混合物を70℃で3時間撹拌した。反応混合物を水(50mL)で希釈し、酢酸エチル(60mL×3)で抽出した。合わせた有機層をブライン(20mL×2)で洗浄した。有機層を無水硫酸ナトリウムで乾燥させ、濾過し、真空下で濃縮して、残渣を得た。残渣を、分取HPLC(カラム:Phenomenex Synergi C18 150*25*10um;移動相:[水(0.225%FA)-ACN];B%:65%~95%、9分)によって精製して、メチル2-[4-クロロ-3-(2,2-ジエトキシエトキシ)イソキサゾール-5-イル]-3-メチル-ブタノエート(337mg、0.965mmol、収率61%)を無色の油として得た。 To a solution of methyl 2-[3-(2,2-diethoxyethoxy)isoxazol-5-yl]-3-methyl-butanoate (500 mg, 1.59 mmol, 1 eq.) in N,N-dimethylformamide (10 mL) was added N-chlorosuccinimide (423 mg, 3.17 mmol, 2 eq.). The reaction mixture was stirred at 70°C for 3 h. The reaction mixture was diluted with water (50 mL) and extracted with ethyl acetate (60 mL x 3). The combined organic layers were washed with brine (20 mL x 2). The organic layers were dried over anhydrous sodium sulfate, filtered, and concentrated under vacuum to give a residue. The residue was purified by preparative HPLC (column: Phenomenex Synergi C18 150*25*10um; mobile phase: [water (0.225% FA)-ACN]; B%: 65%-95%, 9 min) to give methyl 2-[4-chloro-3-(2,2-diethoxyethoxy)isoxazol-5-yl]-3-methyl-butanoate (337 mg, 0.965 mmol, 61% yield) as a colorless oil.
工程2
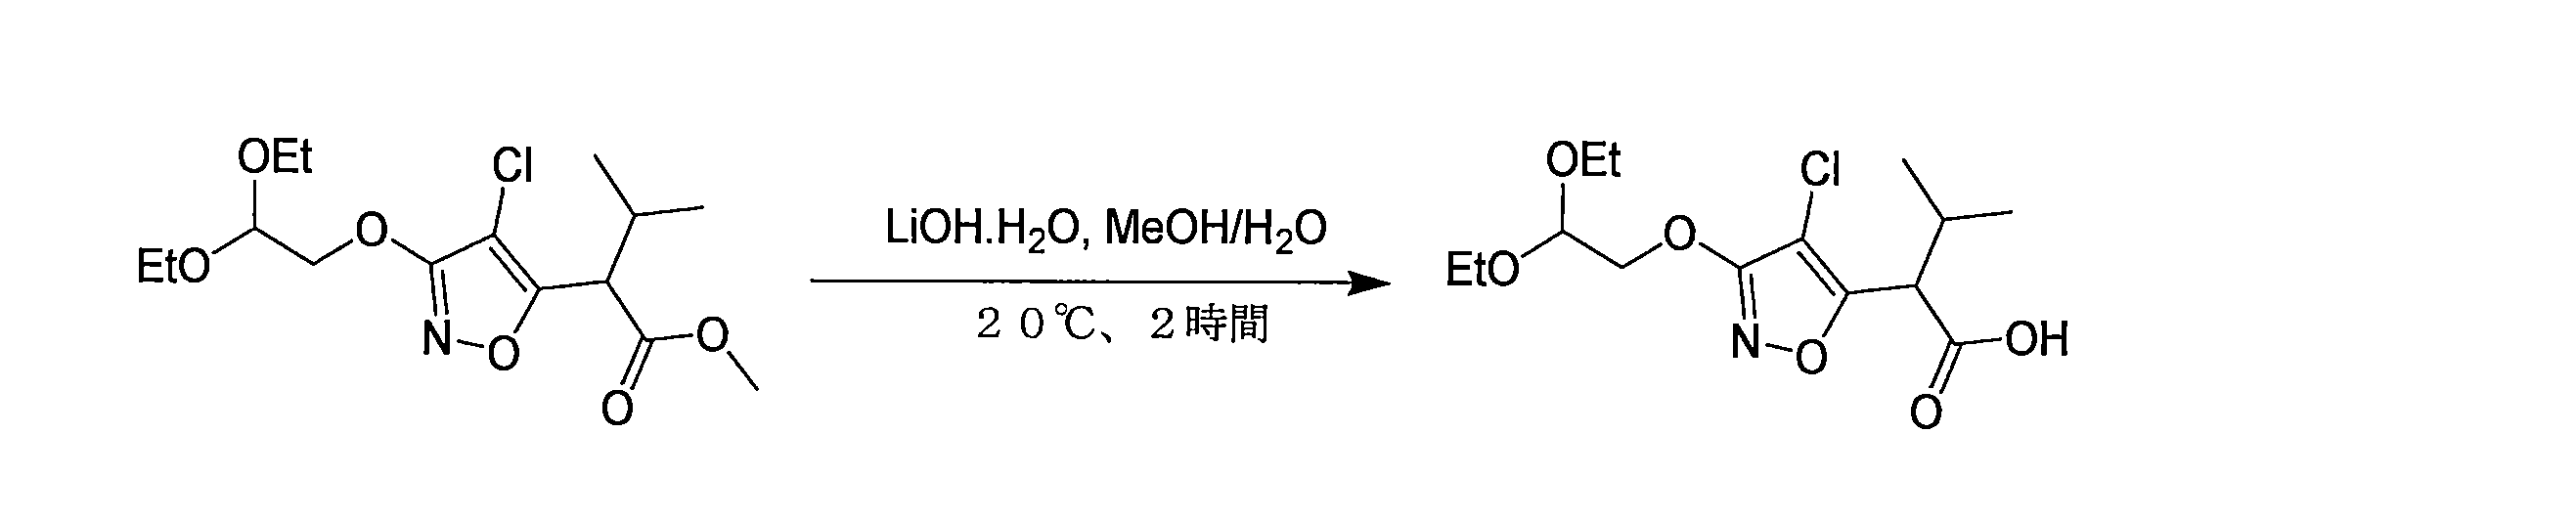
メタノール(2mL)および水(1mL)中のメチル2-[4-クロロ-3-(2,2-ジエトキシエトキシ)イソキサゾール-5-イル]-3-メチル-ブタノエート(377mg、1.08mmol、1当量)の溶液に、水酸化リチウム一水和物(181mg、4.31mmol、4当量)を添加した。反応混合物を20℃で2時間撹拌した。混合物を0℃に冷却し、pHを1M塩化水素で6に調整し、次いで、溶液を真空下で濃縮して、残渣を得た。残渣を、分取HPLC(カラム:Phenomenex luna C18 150*25 10um;移動相:[水(0.225%FA)-ACN];B%:54%~74%、7.8分)によって精製して、生成物を得た。2-[4-クロロ-3-(2,2-ジエトキシエトキシ)イソキサゾール-5-イル]-3-メチル-ブタン酸(229mg、0.68mmol、収率63%)を白色の固体として得た。 To a solution of methyl 2-[4-chloro-3-(2,2-diethoxyethoxy)isoxazol-5-yl]-3-methyl-butanoate (377 mg, 1.08 mmol, 1 eq.) in methanol (2 mL) and water (1 mL) was added lithium hydroxide monohydrate (181 mg, 4.31 mmol, 4 eq.). The reaction mixture was stirred at 20°C for 2 hours. The mixture was cooled to 0°C, the pH was adjusted to 6 with 1M hydrogen chloride, and then the solution was concentrated under vacuum to give a residue. The residue was purified by preparative HPLC (column: Phenomenex luna C18 150*25 10um; mobile phase: [water (0.225% FA)-ACN]; B%: 54%-74%, 7.8 min) to give the product. 2-[4-chloro-3-(2,2-diethoxyethoxy)isoxazol-5-yl]-3-methyl-butanoic acid (229 mg, 0.68 mmol, 63% yield) was obtained as a white solid.
上記の他の実施例について記載される手順を使用して、2-[4-クロロ-3-(2,2-ジエトキシエトキシ)イソキサゾール-5-イル]-3-メチル-ブタン酸を最終化合物に変換した。 2-[4-chloro-3-(2,2-diethoxyethoxy)isoxazol-5-yl]-3-methyl-butanoic acid was converted to the final compound using the procedures described for other examples above.
例示的な化合物222を、類似の手順を使用して調製した。 Exemplary compound 222 was prepared using a similar procedure.
例示的な化合物225の例示的な合成

工程1

四塩化炭素(50mL)中の5-ブロモ-2-メチル-ベンゾニトリル(5g、25.50mmol、1当量)の溶液に、N-ブロモスクシンイミド(4.77g、26.78mmol、1.05当量)および2,2-アゾビス(2-メチルプロピオニトリル)(419mg、2.55mmol、0.1当量)を窒素下で添加した。反応混合物を80℃で16時間撹拌した。混合物を濾過し、濾液を真空中で濃縮した。残渣を、シリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=1:0~10:1)によって精製した。5-ブロモ-2-(ブロモメチル)ベンゾニトリル(4.5g、16.37mmol、収率64%)を淡い黄色の固体として得た。 To a solution of 5-bromo-2-methyl-benzonitrile (5 g, 25.50 mmol, 1 equiv.) in carbon tetrachloride (50 mL), N-bromosuccinimide (4.77 g, 26.78 mmol, 1.05 equiv.) and 2,2-azobis(2-methylpropionitrile) (419 mg, 2.55 mmol, 0.1 equiv.) were added under nitrogen. The reaction mixture was stirred at 80 °C for 16 h. The mixture was filtered and the filtrate was concentrated in vacuum. The residue was purified by silica gel chromatography (petroleum ether: ethyl acetate = 1:0 to 10:1). 5-bromo-2-(bromomethyl)benzonitrile (4.5 g, 16.37 mmol, 64% yield) was obtained as a pale yellow solid.
工程2

アセトニトリル(50mL)中の5-ブロモ-2-(ブロモメチル)ベンゾニトリル(4.4g、16.00mmol、1当量)およびtert-ブチルN-tert-ブトキシカルボニルカルバメート(4.17g、19.20mmol、1.2当量)の溶液に、炭酸カリウム(4.42g、32.01mmol、2当量)を添加した。反応混合物を50℃で12時間撹拌した。LCMSは、5-ブロモ-2-(ブロモメチル)ベンゾニトリルのほとんどが消費されたことを示した。水(100mL)を混合物に加え、得られた混合物を酢酸エチル(50mL×3)で抽出した。合わせた有機相をブライン(50mL)で洗浄し、硫酸ナトリウムで乾燥させ、濾過し、真空中で濃縮した。残渣を、シリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=100:1~5:1)によって精製した。tert-ブチルN-[(4-ブロモ-2-シアノ-フェニル)メチル]-N-tert-ブトキシカルボニル-カルバメート(6.4g、15.56mmol、収率97%)を無色の油として得た。 To a solution of 5-bromo-2-(bromomethyl)benzonitrile (4.4 g, 16.00 mmol, 1 eq.) and tert-butyl N-tert-butoxycarbonylcarbamate (4.17 g, 19.20 mmol, 1.2 eq.) in acetonitrile (50 mL), potassium carbonate (4.42 g, 32.01 mmol, 2 eq.) was added. The reaction mixture was stirred at 50 °C for 12 h. LCMS showed that most of the 5-bromo-2-(bromomethyl)benzonitrile was consumed. Water (100 mL) was added to the mixture, and the resulting mixture was extracted with ethyl acetate (50 mL x 3). The combined organic phase was washed with brine (50 mL), dried over sodium sulfate, filtered, and concentrated in vacuo. The residue was purified by silica gel chromatography (petroleum ether:ethyl acetate = 100:1 to 5:1). tert-Butyl N-[(4-bromo-2-cyano-phenyl)methyl]-N-tert-butoxycarbonyl-carbamate (6.4 g, 15.56 mmol, 97% yield) was obtained as a colorless oil.
工程3

ジオキサン(20mL)中のtert-ブチルN-[(4-ブロモ-2-シアノ-フェニル)メチル]-N-tert-ブトキシカルボニル-カルバメート(2g、4.86mmol、1当量)、4,4,5,5-テトラメチル-2-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)-1,3,2-ジオキサボロラン(1.36g、5.35mmol、1.1当量)の溶液に、酢酸カリウム(1.19g、12.16mmol、2.5当量)および(1,1’-ビス(ジフェニルホスフィノ)フェロセン)パラジウム(II)二塩化物(356mg、0.49mmol、0.1当量)を窒素下で添加した。反応混合物を90℃で12時間撹拌した。混合物を、セライトパッドを通して濾過し、濾液を濃縮して、tert-ブチルN-tert-ブトキシカルボニル-N-[[2-シアノ-4-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)フェニル]メチル]カルバメート(2.2g、粗製)を黒色の油として得た。 To a solution of tert-butyl N-[(4-bromo-2-cyano-phenyl)methyl]-N-tert-butoxycarbonyl-carbamate (2 g, 4.86 mmol, 1 eq.), 4,4,5,5-tetramethyl-2-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)-1,3,2-dioxaborolane (1.36 g, 5.35 mmol, 1.1 eq.) in dioxane (20 mL), potassium acetate (1.19 g, 12.16 mmol, 2.5 eq.) and (1,1'-bis(diphenylphosphino)ferrocene)palladium(II) dichloride (356 mg, 0.49 mmol, 0.1 eq.) were added under nitrogen. The reaction mixture was stirred at 90 °C for 12 h. The mixture was filtered through a pad of Celite, and the filtrate was concentrated to give tert-butyl N-tert-butoxycarbonyl-N-[[2-cyano-4-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)phenyl]methyl]carbamate (2.2 g, crude) as a black oil.
工程4

ジオキサン(20mL)および水(4mL)中のtert-ブチルN-tert-ブトキシカルボニル-N-[[2-シアノ-4-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)フェニル]メチル]カルバメート(2.2g、4.80mmol、1当量)、5-ブロモ-4-メチル-チアゾール(940mg、5.28mmol、1.1当量)の溶液に、炭酸カリウム(1.66g、12.00mmol、2.5当量)および(1,1’-ビス(ジフェニルホスフィノ)フェロセン)パラジウム(II)二塩化物(351mg、0.48mmol、0.1当量)を窒素下で添加した。反応混合物を100℃で12時間撹拌した。水(100mL)を混合物に添加し、得られた混合物を酢酸エチル(50mL×3)で抽出した。合わせた有機相をブライン(50mL)で洗浄し、硫酸ナトリウムで乾燥させ、濾過し、真空中で濃縮した。残渣を、シリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=100:1~1:1)によって精製した。tert-ブチルN-tert-ブトキシカルボニル-N-[[2-シアノ-4-(4-メチルチアゾール-5-イル)フェニル]メチル]カルバメート(1.7g、3.96mmol、収率82%)を淡い黄色の固体として得た。 To a solution of tert-butyl N-tert-butoxycarbonyl-N-[[2-cyano-4-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)phenyl]methyl]carbamate (2.2 g, 4.80 mmol, 1 eq.), 5-bromo-4-methyl-thiazole (940 mg, 5.28 mmol, 1.1 eq.) in dioxane (20 mL) and water (4 mL), potassium carbonate (1.66 g, 12.00 mmol, 2.5 eq.) and (1,1'-bis(diphenylphosphino)ferrocene)palladium(II) dichloride (351 mg, 0.48 mmol, 0.1 eq.) were added under nitrogen. The reaction mixture was stirred at 100 °C for 12 h. Water (100 mL) was added to the mixture and the resulting mixture was extracted with ethyl acetate (50 mL x 3). The combined organic phase was washed with brine (50 mL), dried over sodium sulfate, filtered, and concentrated in vacuo. The residue was purified by silica gel chromatography (petroleum ether:ethyl acetate=100:1 to 1:1). tert-Butyl N-tert-butoxycarbonyl-N-[[2-cyano-4-(4-methylthiazol-5-yl)phenyl]methyl]carbamate (1.7 g, 3.96 mmol, 82% yield) was obtained as a pale yellow solid.
他の実施例について上に記載されるように、tert-ブチルN-tert-ブトキシカルボニル-N-[[2-シアノ-4-(4-メチルチアゾール-5-イル)フェニル]メチル]カルバメートを最終化合物に変換した。 tert-Butyl N-tert-butoxycarbonyl-N-[[2-cyano-4-(4-methylthiazol-5-yl)phenyl]methyl]carbamate was converted to the final compound as described above for other examples.
例示的な化合物224を、類似の手順を使用して調製した。 Exemplary compound 224 was prepared using a similar procedure.
例示的な化合物126の例示的な合成
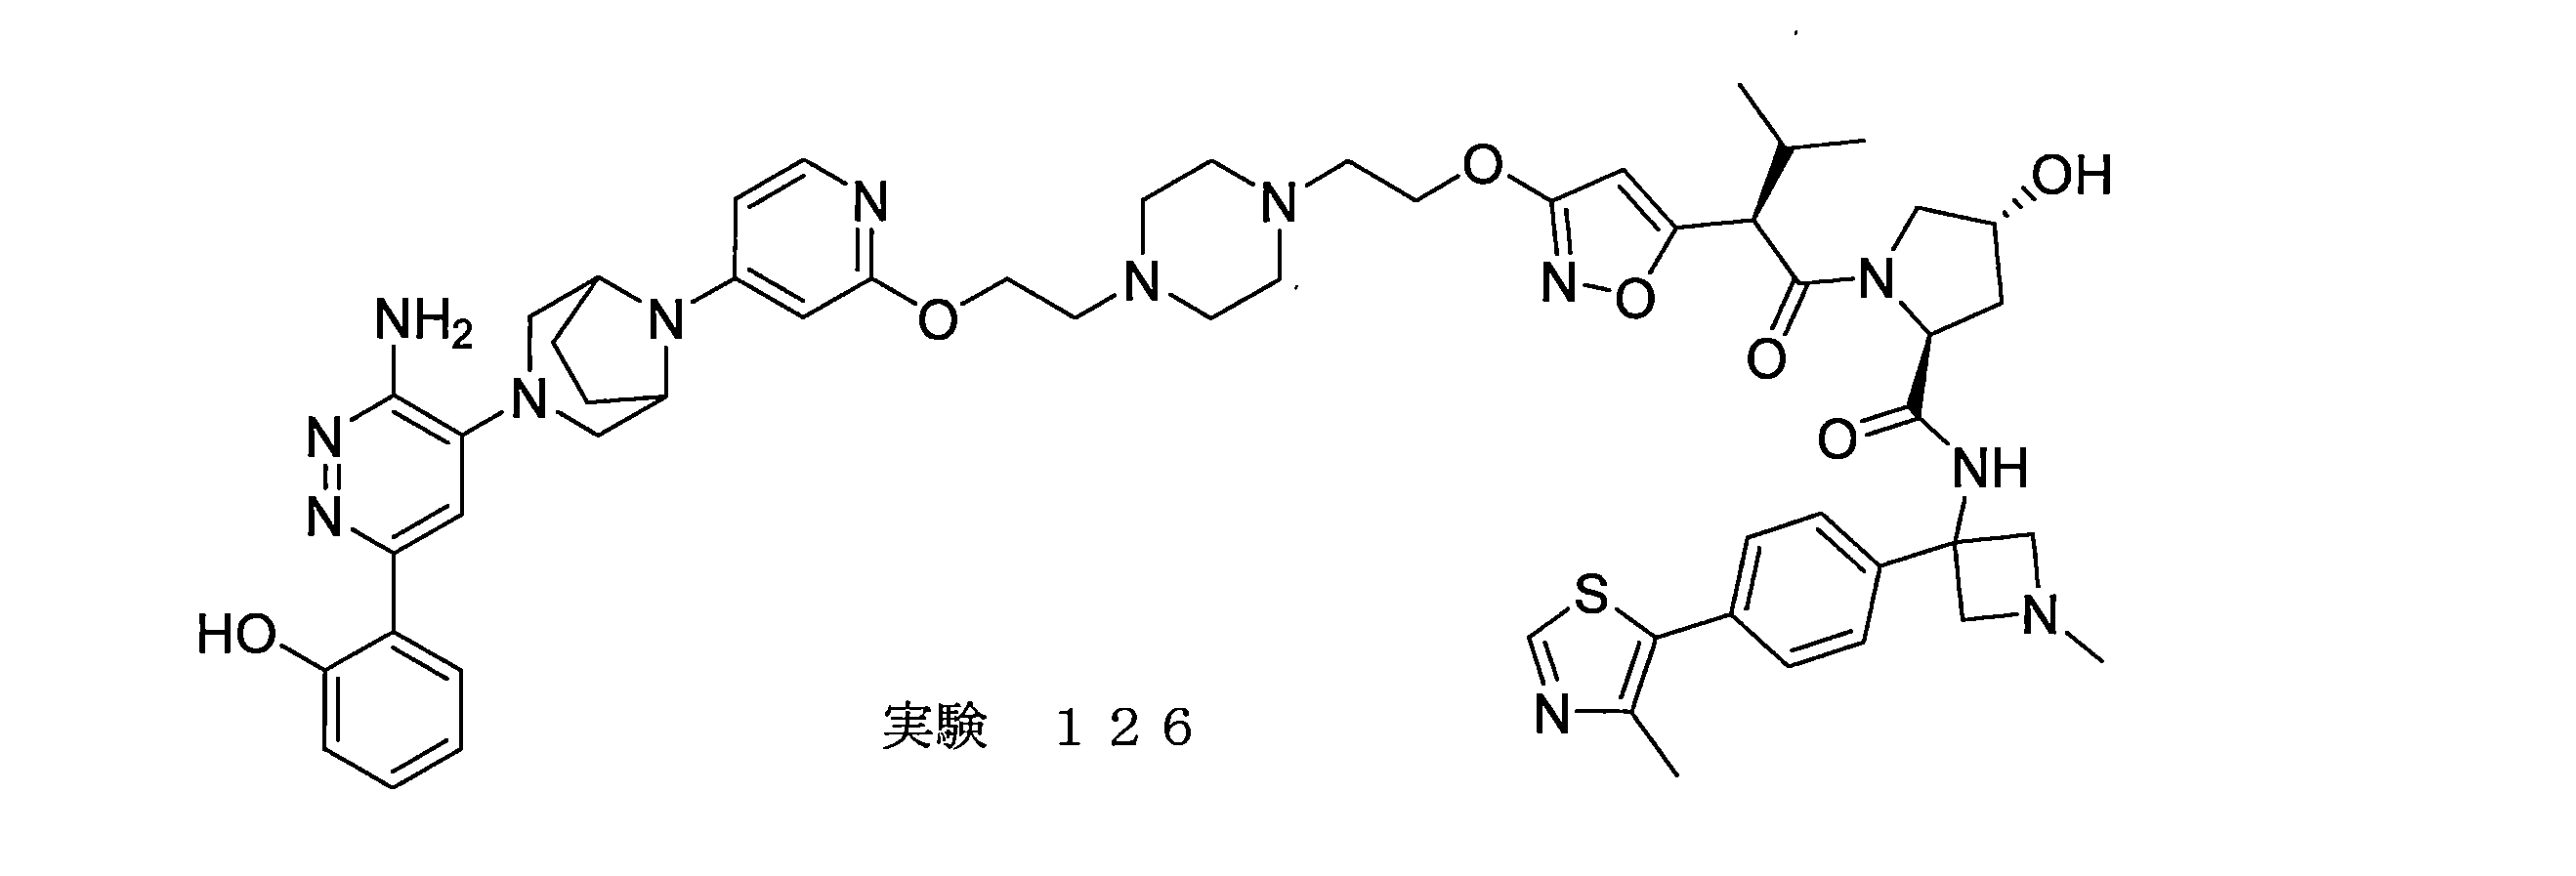
工程1

THF(250mL)中のベンジル3-オキソアゼチジン-1-カルボキシレート(25g、122mmol、1当量)および2-メチルプロパン-2-スルフィンアミド(14.8g、122mmol、1当量)の混合物に、チタン(IV)イソプロピラート(38.9g、171mmol、35.4mL、1.4当量)を窒素下で、15℃で一度に添加した。混合物を60℃で2時間撹拌した。混合物を、酢酸エチルおよび氷水(1000mL)に注ぎ入れ、20分間撹拌し、濾過した。水相を酢酸エチル(500mL×3)で抽出した。合わせた有機相をブライン(200mL×3)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、真空中で濃縮した。残渣を、シリカゲルクロマトグラフィー(石油エーテル/酢酸エチル=30/1~8/1)によって精製して、ベンジル3-tert-ブチルスルフィニルイミノアゼチジン-1-カルボキシレート(39g、粗製)を黄色の固体として得た。 To a mixture of benzyl 3-oxoazetidine-1-carboxylate (25 g, 122 mmol, 1 eq.) and 2-methylpropane-2-sulfinamide (14.8 g, 122 mmol, 1 eq.) in THF (250 mL), titanium(IV) isopropylate (38.9 g, 171 mmol, 35.4 mL, 1.4 eq.) was added in one portion under nitrogen at 15 °C. The mixture was stirred at 60 °C for 2 h. The mixture was poured into ethyl acetate and ice water (1000 mL), stirred for 20 min, and filtered. The aqueous phase was extracted with ethyl acetate (500 mL x 3). The combined organic phase was washed with brine (200 mL x 3), dried over anhydrous sodium sulfate, filtered, and concentrated in vacuum. The residue was purified by silica gel chromatography (petroleum ether/ethyl acetate = 30/1 to 8/1) to give benzyl 3-tert-butylsulfinyliminoazetidine-1-carboxylate (39 g, crude) as a yellow solid.
工程2

THF(350mL)中の1-ブロモ-4-ヨード-ベンゼン(35.78g、126mmol、1当量)の溶液に、n-ブチルリチウム(2.5M、50.6mL、1当量)を窒素下で、-70℃で、滴下で添加し、混合物をこの温度で30分間撹拌した。次いで、THF(70mL)中のベンジル3-tert-ブチルスルフィニルイミノアゼチジン-1-カルボキシレート(39g、126.46mmol、1当量)の溶液を窒素下で、滴下で添加した。反応混合物を-70℃で1時間撹拌した。混合物を氷水(200mL)でクエンチした。水相を酢酸エチル(500mL×3)で抽出した。合わせた有機相をブライン(200mL×2)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、真空中で濃縮した。残渣を、シリカゲルクロマトグラフィー(石油エーテル/酢酸エチル=3/1~1/1)によって精製した。次いで、粗生成物を、半分取逆相HPLC(カラム:Phenomenex luna c18 250mm*100mm*10um;移動相:[水(0.1%TFA)-ACN];B%:40ACN%~70ACN%、45分)によって精製して、ベンジル3-(4-ブロモフェニル)-3-(tert-ブチルスルフィニルアミノ)アゼチジン-1-カルボキシレート(4.62g、9.93mmol、収率7%)を黄色の油として得た。加えて、4.16gのやや不純な生成物(純度86%)を黄色の油として回収した。 To a solution of 1-bromo-4-iodo-benzene (35.78 g, 126 mmol, 1 equiv.) in THF (350 mL), n-butyllithium (2.5 M, 50.6 mL, 1 equiv.) was added dropwise under nitrogen at −70° C., and the mixture was stirred at this temperature for 30 min. Then, a solution of benzyl 3-tert-butylsulfinyl iminoazetidine-1-carboxylate (39 g, 126.46 mmol, 1 equiv.) in THF (70 mL) was added dropwise under nitrogen. The reaction mixture was stirred at −70° C. for 1 h. The mixture was quenched with ice water (200 mL). The aqueous phase was extracted with ethyl acetate (500 mL × 3). The combined organic phase was washed with brine (200 mL × 2), dried over anhydrous sodium sulfate, filtered, and concentrated in vacuum. The residue was purified by silica gel chromatography (petroleum ether/ethyl acetate = 3/1 to 1/1). The crude product was then purified by semi-preparative reverse phase HPLC (column: Phenomenex luna c18 250mm*100mm*10um; mobile phase: [water (0.1% TFA)-ACN]; B%: 40ACN% to 70ACN%, 45min) to give benzyl 3-(4-bromophenyl)-3-(tert-butylsulfinylamino)azetidine-1-carboxylate (4.62g, 9.93mmol, 7% yield) as a yellow oil. Additionally, 4.16g of slightly impure product (86% purity) was recovered as a yellow oil.
工程3

ジメチルアセトアミド(60mL)中のベンジル3-(4-ブロモフェニル)-3-(tert-ブチルスルフィニルアミノ)アゼチジン-1-カルボキシレート(4.62g、9.93mmol、1当量)、酢酸カリウム(1.95g、19.85mmol、2当量)、および4-メチルチアゾール(1.97g、19.85mmol、1.8mL、2当量)の混合物に、酢酸パラジウム(II)(223mg、0.99mmol、0.1当量)を窒素下で、20℃で一度に添加した。混合物を100℃で16時間撹拌した。混合物を20℃に冷却し、氷水(w/w=1/1、100mL)に注ぎ入れ、5分間撹拌した。水相を酢酸エチル(50mL×4)で抽出した。合わせた有機相をブライン(30mL×2)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、真空中で濃縮した。残渣を、半分取逆相HPLC(ギ酸条件;カラム:Phenomenex Synergi Max-RP 250*50mm*10um;移動相:[水(0.225%FA)-ACN];B%:45ACN%~75ACN%、29分)によって精製して、ベンジル3-(tert-ブチルスルフィニルアミノ)-3-[4-(4-メチルチアゾール-5-イル)フェニル]アゼチジン-1-カルボキシレート(2.3g、4.76mmol、収率48%)を黄色の固体として得た。 To a mixture of benzyl 3-(4-bromophenyl)-3-(tert-butylsulfinylamino)azetidine-1-carboxylate (4.62 g, 9.93 mmol, 1 eq.), potassium acetate (1.95 g, 19.85 mmol, 2 eq.), and 4-methylthiazole (1.97 g, 19.85 mmol, 1.8 mL, 2 eq.) in dimethylacetamide (60 mL), palladium(II) acetate (223 mg, 0.99 mmol, 0.1 eq.) was added in one portion at 20°C under nitrogen. The mixture was stirred at 100°C for 16 hours. The mixture was cooled to 20°C, poured into ice water (w/w=1/1, 100 mL), and stirred for 5 minutes. The aqueous phase was extracted with ethyl acetate (50 mL x 4). The combined organic phase was washed with brine (30 mL x 2), dried over anhydrous sodium sulfate, filtered, and concentrated in vacuo. The residue was purified by semi-preparative reverse phase HPLC (formic acid conditions; column: Phenomenex Synergi Max-RP 250*50mm*10um; mobile phase: [water (0.225% FA)-ACN]; B%: 45ACN% to 75ACN%, 29 min) to give benzyl 3-(tert-butylsulfinylamino)-3-[4-(4-methylthiazol-5-yl)phenyl]azetidine-1-carboxylate (2.3 g, 4.76 mmol, 48% yield) as a yellow solid.
工程4

ジクロロメタン(5mL)中のベンジル3-(tert-ブチルスルフィニルアミノ)-3-[4-(4-メチルチアゾール-5-イル)フェニル]アゼチジン-1-カルボキシレート(3.4g、7.03mmol、1当量)および塩酸/メタノール(4M、30mL、17.07当量)の溶液を、25℃で2時間撹拌した。化合物ベンジル3-アミノ-3-[4-(4-メチルチアゾール-5-イル)フェニル]アゼチジン-1-カルボキシレート塩酸塩(3g、粗製)を白色の固体として得た。 A solution of benzyl 3-(tert-butylsulfinylamino)-3-[4-(4-methylthiazol-5-yl)phenyl]azetidine-1-carboxylate (3.4 g, 7.03 mmol, 1 eq.) and hydrochloric acid/methanol (4 M, 30 mL, 17.07 eq.) in dichloromethane (5 mL) was stirred at 25°C for 2 h. The compound benzyl 3-amino-3-[4-(4-methylthiazol-5-yl)phenyl]azetidine-1-carboxylate hydrochloride (3 g, crude) was obtained as a white solid.
工程5

N,N-ジメチルホルムアミド(40mL)中のベンジル3-アミノ-3-[4-(4-メチルチアゾール-5-イル)フェニル]アゼチジン-1-カルボキシレート塩酸塩(3g、7.21mmol、1当量)、O-(7-アザベンゾトリアゾール-1-イル)-N,N,N’,N’-テトラメチルウロニウムヘキサフルオロホスフェート(4.11g、10.82mmol、1.5当量)、および(2S,4R)-1-tert-ブトキシカルボニル-4-ヒドロキシ-ピロリジン-2-カルボン酸(1.67g、7.21mmol、1当量)の混合物に、トリエチルアミン(2.19g、21.64mmol、3.0mL、3当量)を窒素下で、15℃で一度に添加した。混合物を15℃で1時間撹拌した。混合物を氷水(w/w=1/1、100mL)に注ぎ入れ、5分間撹拌した。水相を酢酸エチル(60mL×3)で抽出した。合わせた有機相をブライン(30mL×2)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、真空下で濃縮した。残渣を、半分取逆相HPLC(カラム:Phenomenex Synergi Max-RP 250*80mm*10um;移動相:[水(0.225%FA)-ACN];B%:30%~60%、30分)によって精製した。化合物tert-ブチル(2S,4R)-2-[[1-ベンジルオキシカルボニル-3-[4-(4-メチルチアゾール-5-イル)フェニル]アゼチジン-3-イル]カルバモイル]-4-ヒドロキシ-ピロリジン-1-カルボキシレート(3.32g、5.60mmol、収率77%)を白色の固体として得た。 To a mixture of benzyl 3-amino-3-[4-(4-methylthiazol-5-yl)phenyl]azetidine-1-carboxylate hydrochloride (3 g, 7.21 mmol, 1 eq.), O-(7-azabenzotriazol-1-yl)-N,N,N',N'-tetramethyluronium hexafluorophosphate (4.11 g, 10.82 mmol, 1.5 eq.), and (2S,4R)-1-tert-butoxycarbonyl-4-hydroxy-pyrrolidine-2-carboxylic acid (1.67 g, 7.21 mmol, 1 eq.) in N,N-dimethylformamide (40 mL), triethylamine (2.19 g, 21.64 mmol, 3.0 mL, 3 eq.) was added in one portion under nitrogen at 15°C. The mixture was stirred at 15°C for 1 h. The mixture was poured into ice water (w/w=1/1, 100 mL) and stirred for 5 min. The aqueous phase was extracted with ethyl acetate (60 mL×3). The combined organic phase was washed with brine (30 mL×2), dried over anhydrous sodium sulfate, filtered and concentrated under vacuum. The residue was purified by semi-preparative reverse phase HPLC (column: Phenomenex Synergi Max-RP 250*80 mm*10 um; mobile phase: [water (0.225% FA)-ACN]; B%: 30%-60%, 30 min). The compound tert-butyl (2S,4R)-2-[[1-benzyloxycarbonyl-3-[4-(4-methylthiazol-5-yl)phenyl]azetidin-3-yl]carbamoyl]-4-hydroxy-pyrrolidine-1-carboxylate (3.32 g, 5.60 mmol, 77% yield) was obtained as a white solid.
工程6

ジクロロメタン(100mL)中のtert-ブチル(2S,4R)-2-[[1-ベンジルオキシカルボニル-3-[4-(4-メチルチアゾール-5-イル)フェニル]アゼチジン-3-イル]カルバモイル]-4-ヒドロキシ-ピロリジン-1-カルボキシレート(1.1g、1.86mmol、1当量)およびトリエチルアミン(666mg、6.59mmol、0.9mL、3.55当量)の混合物に、塩化パラジウム(ii)(329.10mg、1.86mmol、1当量)を窒素下で、0℃で一度に添加した。トリエチルシラン(2.81g、24.13mmol、3.8mL、13当量)を添加し、反応混合物を20℃で2時間撹拌した。反応物に、トリフルオロ酢酸(4.23g、37.14mmol、2.75mL、20.01当量)を添加し、混合物を20℃でさらに1時間撹拌した。混合物を濾過し、40℃において減圧下で濃縮した。残渣を、半分取逆相HPLC(ホルム酸条件;カラム:Kromasil 250*50mm*10um;移動相:[水(0.225%FA)-ACN];B%:26ACN%~56ACN%、28分)によって精製した。tert-ブチル(2S,4R)-4-ヒドロキシ-2-[[3-[4-(4-メチルチアゾール-5-イル)フェニル]アゼチジン-3-イル]カルバモイル]ピロリジン-1-カルボキシレート(336mg、0.73mmol、収率39%)を白色の固体として得た。 To a mixture of tert-butyl (2S,4R)-2-[[1-benzyloxycarbonyl-3-[4-(4-methylthiazol-5-yl)phenyl]azetidin-3-yl]carbamoyl]-4-hydroxy-pyrrolidine-1-carboxylate (1.1 g, 1.86 mmol, 1 equiv.) and triethylamine (666 mg, 6.59 mmol, 0.9 mL, 3.55 equiv.) in dichloromethane (100 mL), palladium(ii) chloride (329.10 mg, 1.86 mmol, 1 equiv.) was added in one portion under nitrogen at 0° C. Triethylsilane (2.81 g, 24.13 mmol, 3.8 mL, 13 equiv.) was added and the reaction mixture was stirred at 20° C. for 2 h. To the reaction, trifluoroacetic acid (4.23 g, 37.14 mmol, 2.75 mL, 20.01 equiv.) was added and the mixture was stirred at 20° C. for an additional hour. The mixture was filtered and concentrated under reduced pressure at 40° C. The residue was purified by semi-preparative reverse phase HPLC (formic acid conditions; column: Kromasil 250*50 mm*10 um; mobile phase: [water (0.225% FA)-ACN]; B%: 26 ACN% to 56 ACN%, 28 min). tert-Butyl (2S,4R)-4-hydroxy-2-[[3-[4-(4-methylthiazol-5-yl)phenyl]azetidin-3-yl]carbamoyl]pyrrolidine-1-carboxylate (336 mg, 0.73 mmol, 39% yield) was obtained as a white solid.
工程7

ジクロロメタン(15mL)中のtert-ブチル(2S,4R)-4-ヒドロキシ-2-[[3-[4-(4-メチルチアゾール-5-イル)フェニル]アゼチジン-3-イル]カルバモイル]ピロリジン-1-カルボキシレート(390mg、0.85mmol、1当量)およびホルムアルデヒド(138mg、1.70mmol、0.1mL、純度37%、2当量)の混合物を、20℃で30分間撹拌した。酢酸(51mg、0.85mmol、1当量)およびトリアセトキシホウ素水素化ナトリウム(901mg、4.25mmol、5当量)を添加し、混合物を20℃で5時間撹拌した。混合物を40℃において減圧下で濃縮した。残渣を、半分取逆相HPLC(ホルム酸条件;カラム:Phenomenex Synergi C18 150*25*10um;移動相:[水(0.225%FA)-ACN];B%:0%~30%、10分)によって精製した。tert-ブチル(2S,4R)-4-ヒドロキシ-2-[[1-メチル-3-[4-(4-メチルチアゾール-5-イル)フェニル]アゼチジン-3-イル]カルバモイル]ピロリジン-1-カルボキシレート(300mg、0.63mmol、収率74%)を黄色の油として得た。 A mixture of tert-butyl (2S,4R)-4-hydroxy-2-[[3-[4-(4-methylthiazol-5-yl)phenyl]azetidin-3-yl]carbamoyl]pyrrolidine-1-carboxylate (390 mg, 0.85 mmol, 1 eq.) and formaldehyde (138 mg, 1.70 mmol, 0.1 mL, 37% purity, 2 eq.) in dichloromethane (15 mL) was stirred at 20°C for 30 min. Acetic acid (51 mg, 0.85 mmol, 1 eq.) and sodium triacetoxyborohydride (901 mg, 4.25 mmol, 5 eq.) were added and the mixture was stirred at 20°C for 5 h. The mixture was concentrated under reduced pressure at 40°C. The residue was purified by semi-preparative reverse phase HPLC (formic acid condition; column: Phenomenex Synergi C18 150*25*10um; mobile phase: [water (0.225% FA)-ACN]; B%: 0%-30%, 10 min). tert-Butyl (2S,4R)-4-hydroxy-2-[[1-methyl-3-[4-(4-methylthiazol-5-yl)phenyl]azetidin-3-yl]carbamoyl]pyrrolidine-1-carboxylate (300 mg, 0.63 mmol, 74% yield) was obtained as a yellow oil.
上記の他の実施例に記載されるものと類似の手順を使用して、tert-ブチル(2S,4R)-4-ヒドロキシ-2-[[1-メチル-3-[4-(4-メチルチアゾール-5-イル)フェニル]アゼチジン-3-イル]カルバモイル]ピロリジン-1-カルボキシレートを最終化合物に変換した。 Using procedures similar to those described in other examples above, tert-butyl (2S,4R)-4-hydroxy-2-[[1-methyl-3-[4-(4-methylthiazol-5-yl)phenyl]azetidin-3-yl]carbamoyl]pyrrolidine-1-carboxylate was converted to the final compound.
例示的な化合物228を、類似の手順を使用して調製した。 Exemplary compound 228 was prepared using a similar procedure.
例示的な化合物233および234の例示的な合成

例示的化合物76および70について記載されるものと類似の手順を使用して、以下のスキームに従って調製した。

例示的な化合物127の例示的な合成

工程1
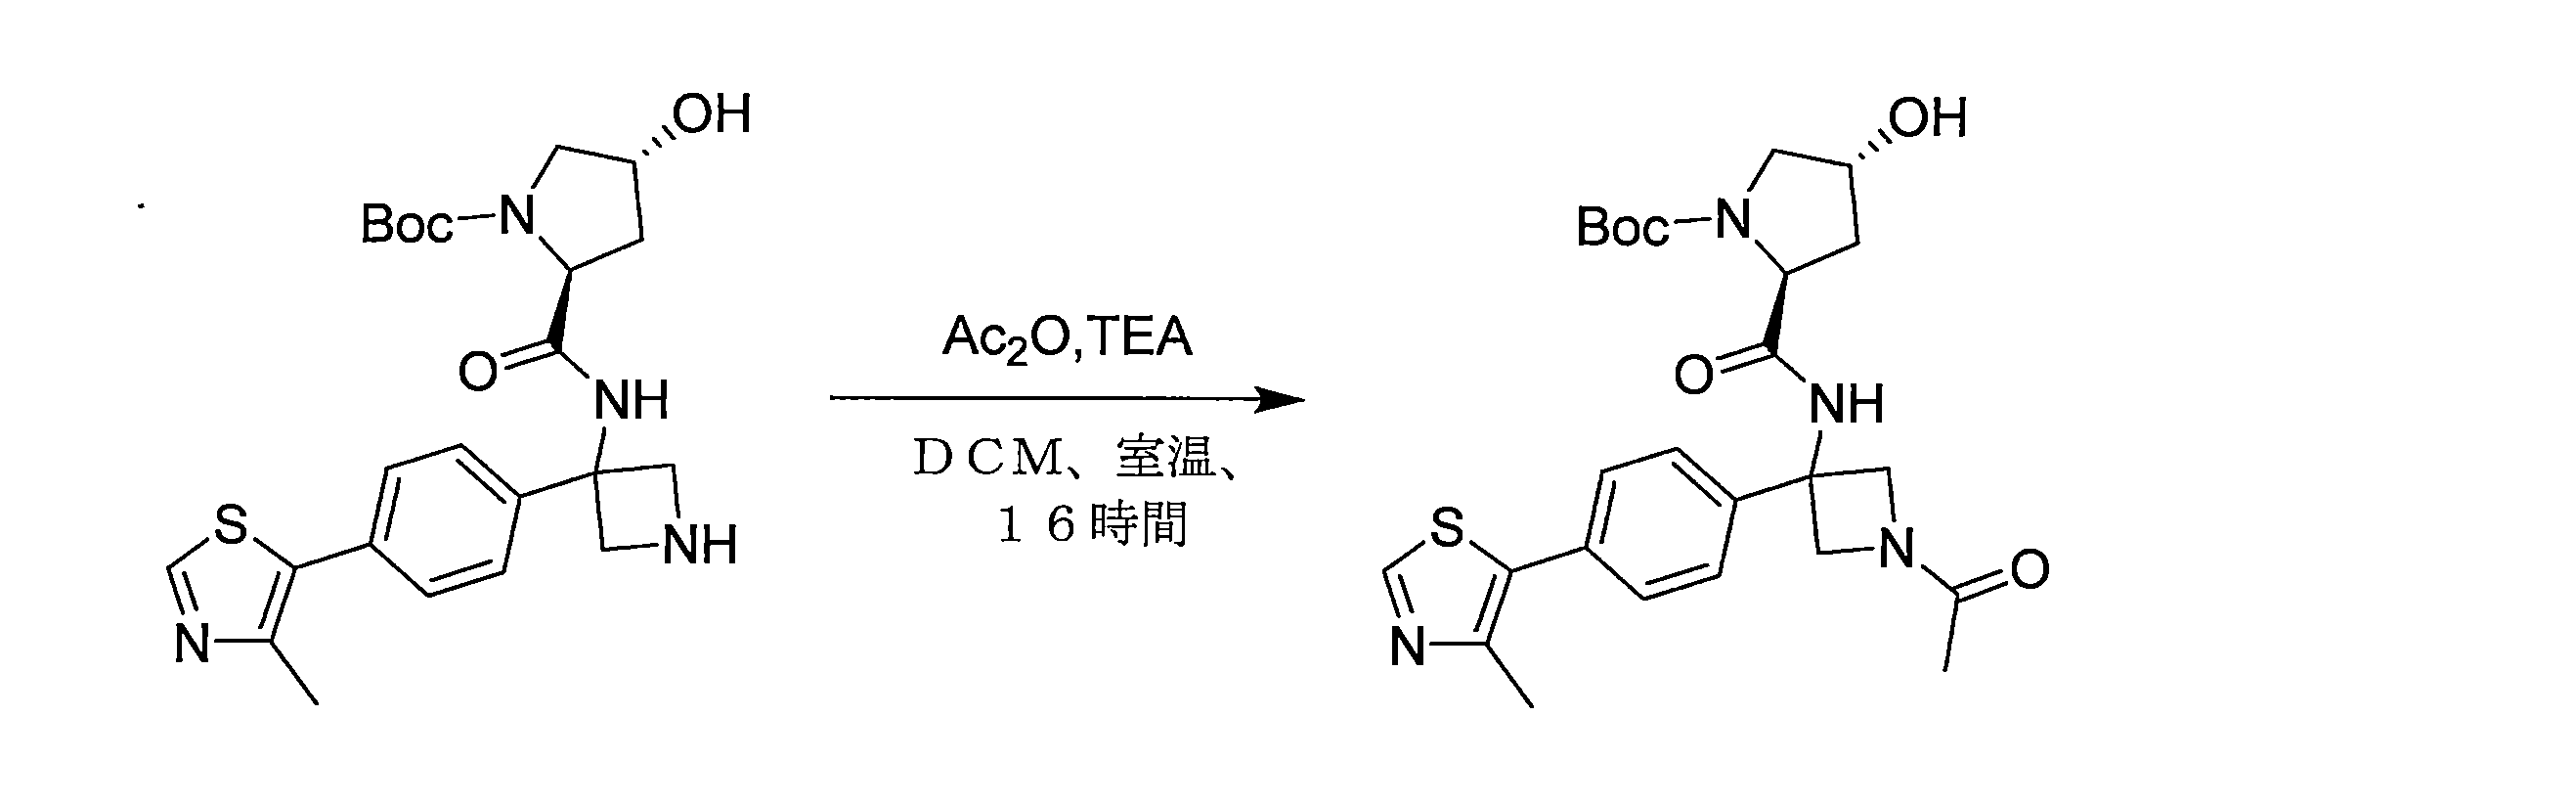
ジクロロメタン(60mL)中のtert-ブチル(2S,4R)-4-ヒドロキシ-2-[[3-[4-(4-メチルチアゾール-5-イル)フェニル]アゼチジン-3-イル]カルバモイル]ピロリジン-1-カルボキシレート(300mg、0.65mmol、1当量)およびトリエチルアミン(198mg、1.96mmol、3当量)の混合物に、無水酢酸(100.18mg、0.98mmol、91.9uL、1.5当量)を窒素下で、25℃で一度に添加した。混合物を25℃で16時間撹拌した。混合物を45℃において減圧下で濃縮した。残渣を、半分取逆相HPLC(基本条件、カラム:Waters Xbridge 150*25 5um;移動相:[水(0.05%水酸化アンモニアv/v)-ACN];B%:10%~40%、10分)によって精製した。粗生成物tert-ブチル(2S,4R)-2-[[1-アセチル-3-[4-(4-メチルチアゾール-5-イル)フェニル]アゼチジン-3-イル]カルバモイル]-4-ヒドロキシ-ピロリジン-1-カルボキシレート(232mg、0.46mmol、収率71%)を黄色の油として得た。 To a mixture of tert-butyl (2S,4R)-4-hydroxy-2-[[3-[4-(4-methylthiazol-5-yl)phenyl]azetidin-3-yl]carbamoyl]pyrrolidine-1-carboxylate (300 mg, 0.65 mmol, 1 eq.) and triethylamine (198 mg, 1.96 mmol, 3 eq.) in dichloromethane (60 mL), acetic anhydride (100.18 mg, 0.98 mmol, 91.9 uL, 1.5 eq.) was added in one portion under nitrogen at 25°C. The mixture was stirred at 25°C for 16 hours. The mixture was concentrated under reduced pressure at 45°C. The residue was purified by semi-preparative reverse phase HPLC (basic conditions, column: Waters Xbridge 150*25 5um; mobile phase: [water (0.05% ammonia hydroxide v/v)-ACN]; B%: 10%-40%, 10 min). The crude product tert-butyl (2S,4R)-2-[[1-acetyl-3-[4-(4-methylthiazol-5-yl)phenyl]azetidin-3-yl]carbamoyl]-4-hydroxy-pyrrolidine-1-carboxylate (232 mg, 0.46 mmol, 71% yield) was obtained as a yellow oil.
上記の実施例について記載される手順を使用して、tert-ブチル(2S,4R)-2-[[1-アセチル-3-[4-(4-メチルチアゾール-5-イル)フェニル]アゼチジン-3-イル]カルバモイル]-4-ヒドロキシ-ピロリジン-1-カルボキシレートを最終生成物に変換した。 Using the procedures described for the examples above, tert-butyl (2S,4R)-2-[[1-acetyl-3-[4-(4-methylthiazol-5-yl)phenyl]azetidin-3-yl]carbamoyl]-4-hydroxy-pyrrolidine-1-carboxylate was converted to the final product.
例示的な化合物235を、類似の手順を使用して調製した。 Exemplary compound 235 was prepared using a similar procedure.
例示的な化合物240の例示的な合成

例示的な化合物202について記載されるものと類似の手順を使用して、以下のスキームに従って、tert-ブチル(S)-(1-(2-フルオロ-4-(4-メチルチアゾール-5-イル)フェニル)エチル)カルバメートを調製した。

他の実施例に記載されるものと類似の手順を使用して、(tert-ブチル(S)-(1-(2-フルオロ-4-(4-メチルチアゾール-5-イル)フェニル)エチル)カルバメートを最終化合物に変換した。 Using procedures similar to those described in other examples, (tert-butyl (S)-(1-(2-fluoro-4-(4-methylthiazol-5-yl)phenyl)ethyl)carbamate was converted to the final compound.
例示的な化合物239を、類似の手順を使用して調製した。 Exemplary compound 239 was prepared using a similar procedure.
例示的な化合物243および244の例示的な合成
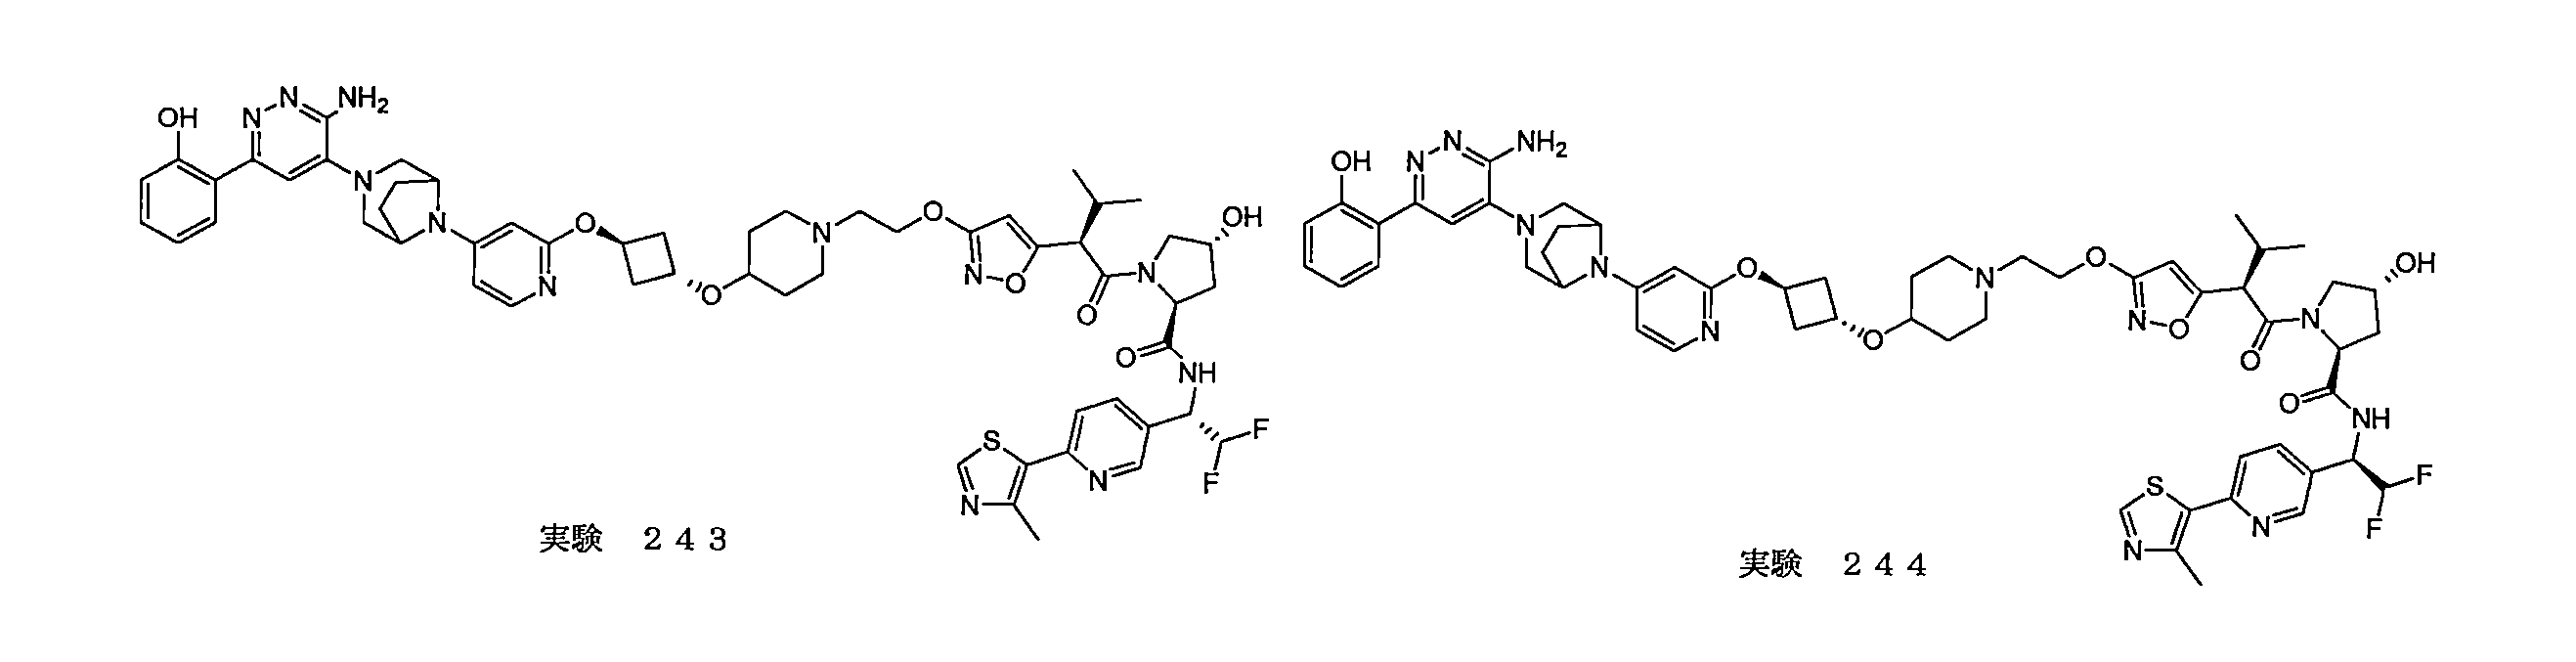
工程1

テトラヒドロフラン(300mL)中のエチルトリフルオロ酢酸塩(21.31g、150mmol、20.7mL、1.5当量)の溶液に、リチウムビス(トリメチルシリル)アミド(1M、120mL、1.2当量)を-78℃で添加した。混合物を-78℃で1時間撹拌した。次いで、テトラヒドロフラン(20mL)中の1-(6-ブロモ-3-ピリジル)エタノン(20g、100mmol、1当量)を、混合物に-78℃で、滴下で添加した。反応混合物を20℃で2時間撹拌した。飽和塩化アンモニウム(200mL)を添加して反応物をクエンチし、水(500mL)を混合物に添加した。得られた混合物を酢酸エチル(300mL×3)で抽出した。合わせた有機相をブライン(500mL)で洗浄し、硫酸ナトリウムで乾燥させ、濾過し、真空中で濃縮した。残渣を、石油エーテルおよび酢酸エチル(255mL、石油エーテル:酢酸エチル=50:1)で粉砕し、濾過し、淡い黄色の固体を得た。濾液を真空中で濃縮した。残渣を、シリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=10:1~0:1)によって精製した。1-(6-ブロモ-3-ピリジル)-4,4,4-トリフルオロ-ブタン-1,3-ジオン(28g、94.6mmol、収率94%)を黄色の固体として得た。 To a solution of ethyl trifluoroacetate (21.31 g, 150 mmol, 20.7 mL, 1.5 eq.) in tetrahydrofuran (300 mL) was added lithium bis(trimethylsilyl)amide (1 M, 120 mL, 1.2 eq.) at −78 °C. The mixture was stirred at −78 °C for 1 h. Then, 1-(6-bromo-3-pyridyl)ethanone (20 g, 100 mmol, 1 eq.) in tetrahydrofuran (20 mL) was added dropwise to the mixture at −78 °C. The reaction mixture was stirred at 20 °C for 2 h. The reaction was quenched by adding saturated ammonium chloride (200 mL) and water (500 mL) was added to the mixture. The resulting mixture was extracted with ethyl acetate (300 mL × 3). The combined organic phase was washed with brine (500 mL), dried over sodium sulfate, filtered, and concentrated in vacuum. The residue was triturated with petroleum ether and ethyl acetate (255 mL, petroleum ether:ethyl acetate=50:1) and filtered to give a pale yellow solid. The filtrate was concentrated in vacuo. The residue was purified by silica gel chromatography (petroleum ether:ethyl acetate=10:1 to 0:1). 1-(6-bromo-3-pyridyl)-4,4,4-trifluoro-butane-1,3-dione (28 g, 94.6 mmol, 94% yield) was obtained as a yellow solid.
工程2

アセトニトリル(150mL)中の1-(6-ブロモ-3-ピリジル)-4,4,4-トリフルオロ-ブタン-1,3-ジオン(15g、50.67mmol、1当量)の溶液に、1-(クロロメチル)-4-フルオロ-1,4-ジアゾニアビシクロ[2.2.2]オクタン;ジテトラフルオロホウ酸塩(44.87g、126.7mmol、2.5当量)を添加した。反応混合物を90℃で3時間撹拌した。次いで、水(1.83g、101.3mmol、1.8mL、2当量)を混合物に添加し、反応混合物を90℃で15分間撹拌した。次いで、トリエチルアミン(25.64g、253.3mmol、35.2mL、5当量)を混合物に20℃で添加し、反応混合物を20℃で16時間撹拌した。水(300mL)を混合物に添加した。得られた混合物を酢酸エチル(300mL×3)で抽出した。合わせた有機相をブライン(500mL)で洗浄し、硫酸ナトリウムで乾燥させ、濾過し、真空中で濃縮した。残渣を、シリカゲルクロマトグラフィー(ジクロロメタン:メタノール=1:0~10:1)によって精製した。1-(6-ブロモ-3-ピリジル)-2,2-ジフルオロ-エタノン(7.9g、粗製)を茶色の油として得た。 To a solution of 1-(6-bromo-3-pyridyl)-4,4,4-trifluoro-butane-1,3-dione (15 g, 50.67 mmol, 1 eq.) in acetonitrile (150 mL), 1-(chloromethyl)-4-fluoro-1,4-diazoniabicyclo[2.2.2]octane; ditetrafluoroborate (44.87 g, 126.7 mmol, 2.5 eq.) was added. The reaction mixture was stirred at 90 °C for 3 h. Then, water (1.83 g, 101.3 mmol, 1.8 mL, 2 eq.) was added to the mixture and the reaction mixture was stirred at 90 °C for 15 min. Then, triethylamine (25.64 g, 253.3 mmol, 35.2 mL, 5 eq.) was added to the mixture at 20 °C and the reaction mixture was stirred at 20 °C for 16 h. Water (300 mL) was added to the mixture. The resulting mixture was extracted with ethyl acetate (300 mL x 3). The combined organic phase was washed with brine (500 mL), dried over sodium sulfate, filtered, and concentrated in vacuo. The residue was purified by silica gel chromatography (dichloromethane:methanol = 1:0 to 10:1). 1-(6-bromo-3-pyridyl)-2,2-difluoro-ethanone (7.9 g, crude) was obtained as a brown oil.
工程3
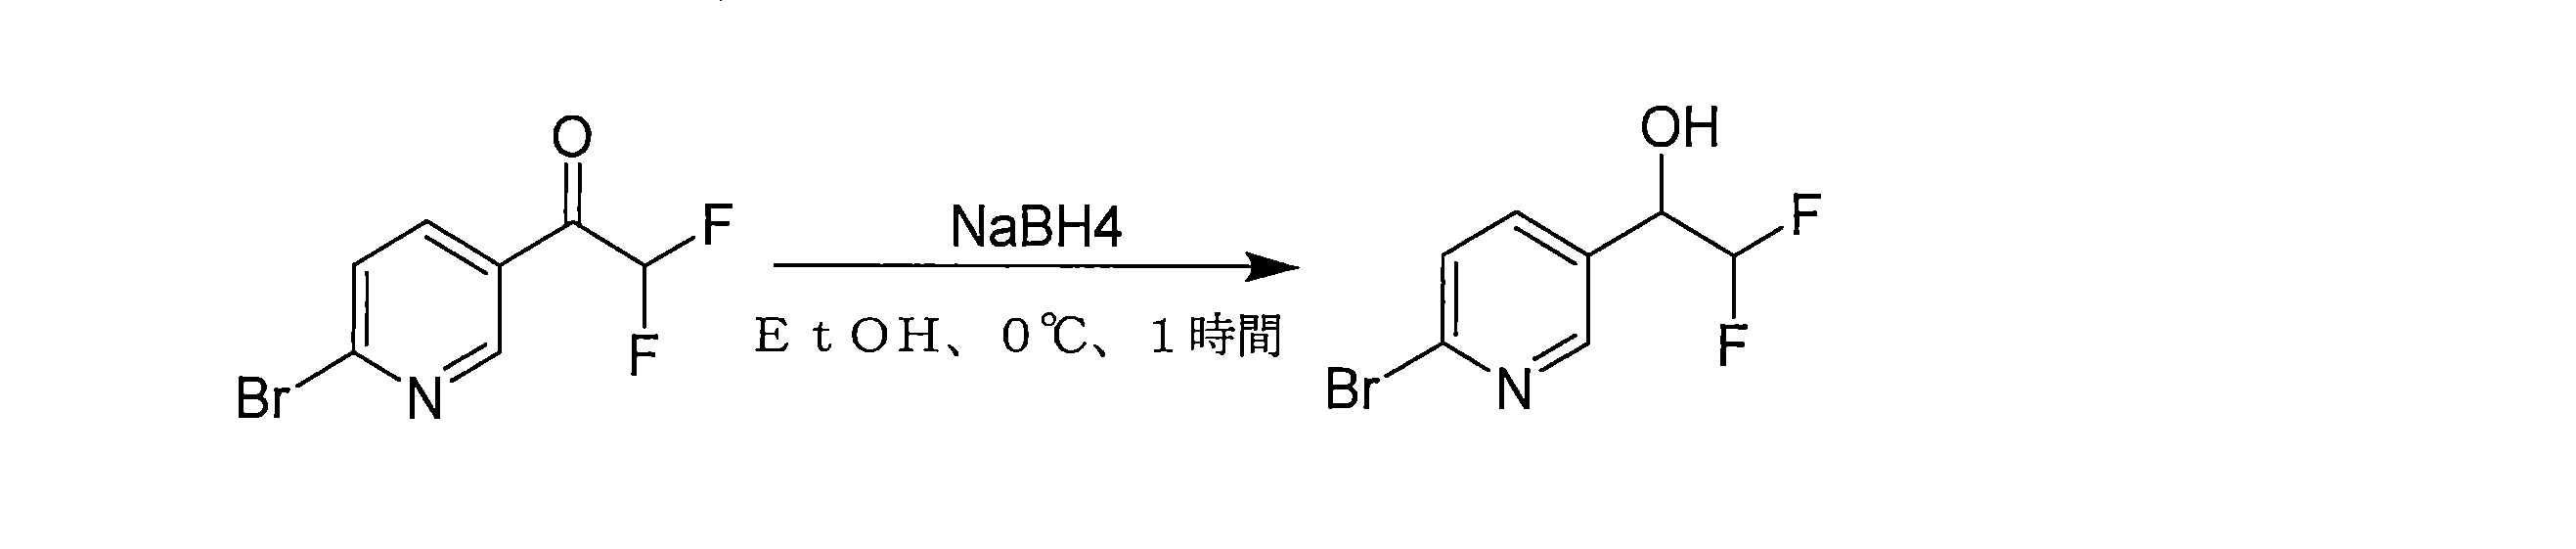
エタノール(80mL)中の1-(6-ブロモ-3-ピリジル)-2,2-ジフルオロ-エタノン(7.9g、33.47mmol、1当量)の溶液に、水素化ホウ素ナトリウム(1.27g、33.47mmol、1当量)を3回に分けて0℃で添加した。反応混合物を0℃で1時間撹拌した。水(200mL)を混合物に添加し、得られた混合物を酢酸エチル(100mL×3)で抽出した。合わせた有機相をブライン(150mL)で洗浄し、硫酸ナトリウムで乾燥させ、濾過し、真空中で濃縮した。残渣を、シリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=100:1~1:1)によって精製した。1-(6-ブロモ-3-ピリジル)-2,2-ジフルオロ-エタノール(5.3g、21.20mmol、収率63%)を淡い黄色の油として得た。 To a solution of 1-(6-bromo-3-pyridyl)-2,2-difluoro-ethanone (7.9 g, 33.47 mmol, 1 equiv.) in ethanol (80 mL) was added sodium borohydride (1.27 g, 33.47 mmol, 1 equiv.) in three portions at 0 °C. The reaction mixture was stirred at 0 °C for 1 h. Water (200 mL) was added to the mixture, and the resulting mixture was extracted with ethyl acetate (100 mL x 3). The combined organic phase was washed with brine (150 mL), dried over sodium sulfate, filtered, and concentrated in vacuum. The residue was purified by silica gel chromatography (petroleum ether: ethyl acetate = 100: 1 to 1: 1). 1-(6-bromo-3-pyridyl)-2,2-difluoro-ethanol (5.3 g, 21.20 mmol, 63% yield) was obtained as a pale yellow oil.
工程4
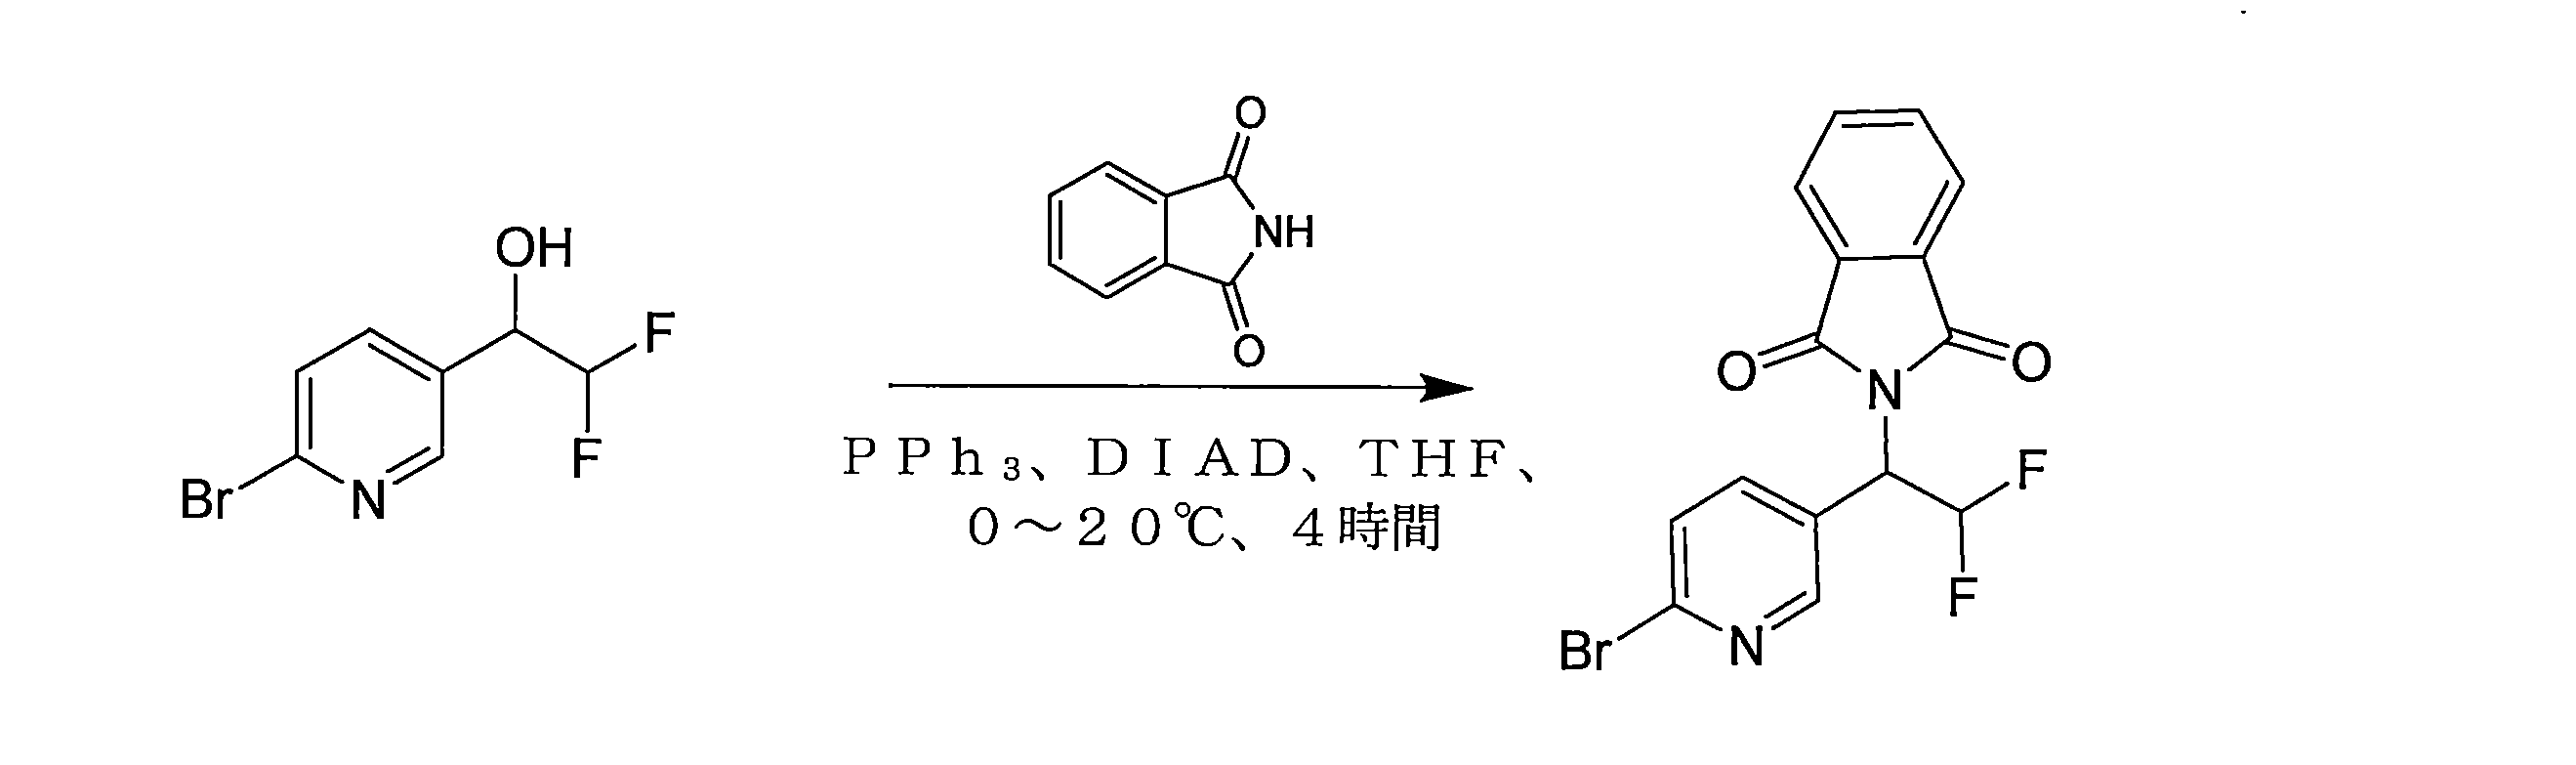
テトラヒドロフラン(30mL)中の1-(6-ブロモ-3-ピリジル)-2,2-ジフルオロ-エタノール(3.3g、13.86mmol、1当量)およびイソインドリン-1,3-ジオン(2.45g、16.64mmol、1.2当量)の溶液に、トリフェニルホスフィン(4.36g、16.64mmol、1.2当量)、続いて、滴下で、テトラヒドロフラン(5mL)中のジイソプロピルアゾジカルボキシレート(3.36g、16.64mmol、3.2mL、1.2当量)を窒素下で、0℃で添加した。反応混合物を20℃で12時間撹拌した。混合物を真空中で濃縮した。残渣を、シリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=1:0~5:1)によって精製した。2-[1-(6-ブロモ-3-ピリジル)-2,2-ジフルオロ-エチル]イソインドリン-1,3-ジオン(2.8g、6.89mmol、収率49%)を淡い黄色の固体として得た。 To a solution of 1-(6-bromo-3-pyridyl)-2,2-difluoro-ethanol (3.3 g, 13.86 mmol, 1 eq.) and isoindoline-1,3-dione (2.45 g, 16.64 mmol, 1.2 eq.) in tetrahydrofuran (30 mL) was added triphenylphosphine (4.36 g, 16.64 mmol, 1.2 eq.) followed by dropwise diisopropyl azodicarboxylate (3.36 g, 16.64 mmol, 3.2 mL, 1.2 eq.) in tetrahydrofuran (5 mL) under nitrogen at 0°C. The reaction mixture was stirred at 20°C for 12 h. The mixture was concentrated in vacuum. The residue was purified by silica gel chromatography (petroleum ether:ethyl acetate=1:0 to 5:1). 2-[1-(6-bromo-3-pyridyl)-2,2-difluoro-ethyl]isoindoline-1,3-dione (2.8 g, 6.89 mmol, 49% yield) was obtained as a pale yellow solid.
工程5

エタノール(20mL)中の2-[1-(6-ブロモ-3-ピリジル)-2,2-ジフルオロ-エチル]イソインドリン-1,3-ジオン(2g、5.45mmol、1当量)の溶液に、ヒドラジン水和物(1.36g、27.24mmol、1.3mL、5当量)を添加した。反応混合物を80℃で2時間撹拌した。エタノール(30mL)を混合物に添加し、混合物を濾過し、濾液を真空中で濃縮した。残渣を次の工程で直接使用した。1-(6-ブロモ-3-ピリジル)-2,2-ジフルオロ-エタンアミン(1.2g、粗製)を淡い黄色の固体として得た。 To a solution of 2-[1-(6-bromo-3-pyridyl)-2,2-difluoro-ethyl]isoindoline-1,3-dione (2 g, 5.45 mmol, 1 eq.) in ethanol (20 mL) was added hydrazine hydrate (1.36 g, 27.24 mmol, 1.3 mL, 5 eq.). The reaction mixture was stirred at 80 °C for 2 h. Ethanol (30 mL) was added to the mixture, the mixture was filtered, and the filtrate was concentrated in vacuo. The residue was used directly in the next step. 1-(6-bromo-3-pyridyl)-2,2-difluoro-ethanamine (1.2 g, crude) was obtained as a pale yellow solid.
工程6
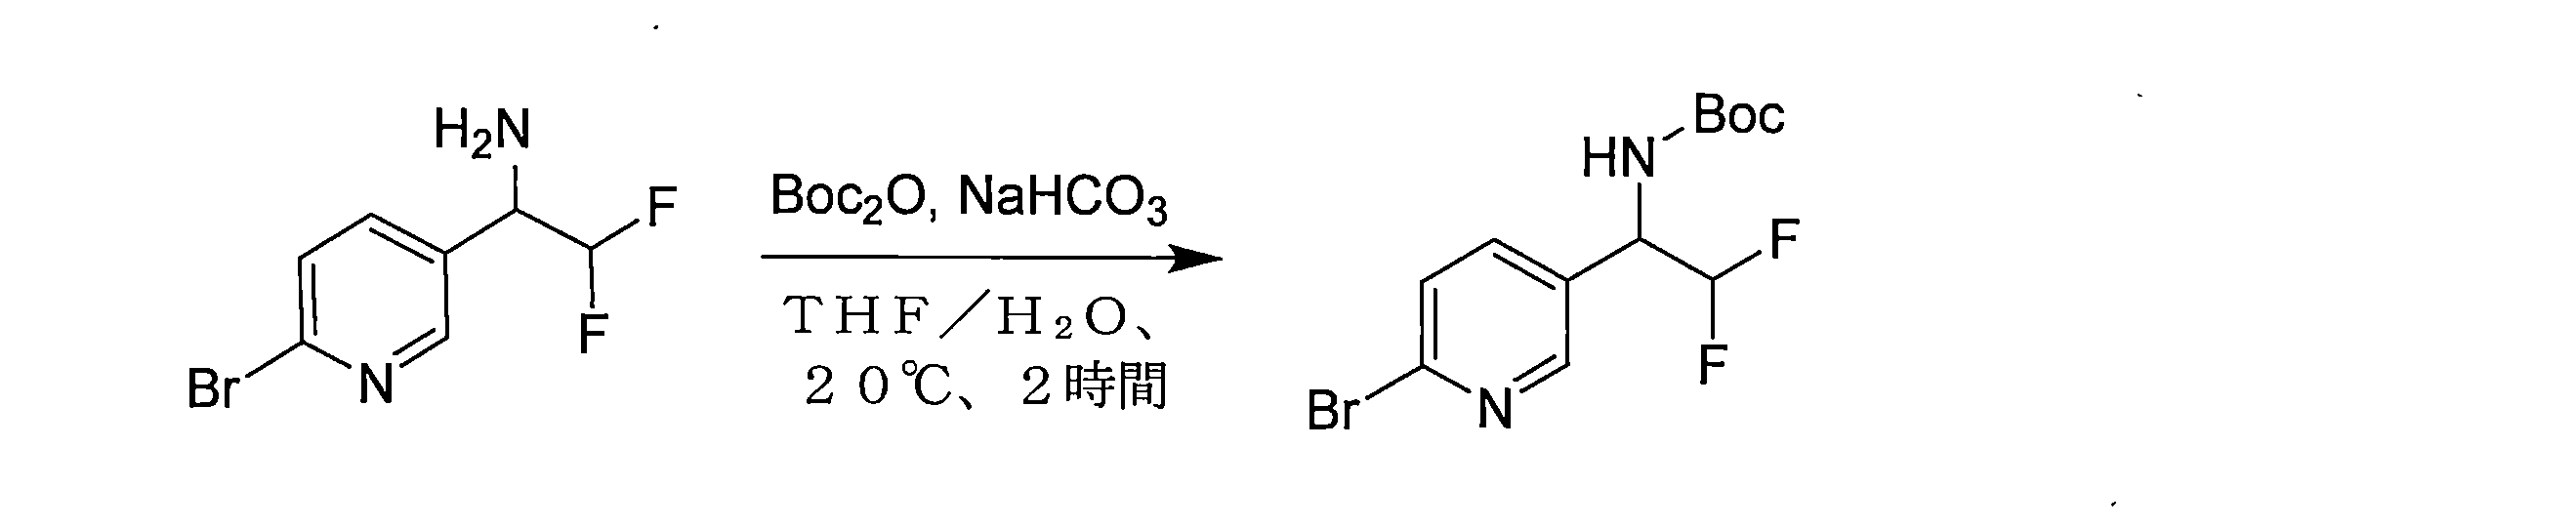
テトラヒドロフラン(5mL)および水(5mL)中の1-(6-ブロモ-3-ピリジル)-2,2-ジフルオロ-エタンアミン(1.2g、5.06mmol、1当量)および重炭酸ナトリウム(850mg、10.12mmol、2当量)の溶液に、ジ-tert-ブチルジカーボネート(2.21g、10.12mmol、2.3mL、2当量)を添加した。反応混合物を20℃で2時間撹拌した。水(100mL)を混合物に加え、得られた混合物を酢酸エチル(50mL×3)で抽出した。合わせた有機相をブライン(60mL)で洗浄し、硫酸ナトリウムで乾燥させ、濾過し、真空中で濃縮した。残渣を、シリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=3:1)によって精製した。粗生成物を、分取高速液体(カラム:Phenomenex Gemini C18 250*50mm*10um;移動相:[水(0.05%水酸化アンモニアv/v)-ACN];B%:40%~65%、22分)によって精製した。tert-ブチルN-[1-(6-ブロモ-3-ピリジル)-2,2-ジフルオロ-エチル]カルバメート(1.4g、4.15mmol、収率82%)を淡い黄色の固体として得た。 To a solution of 1-(6-bromo-3-pyridyl)-2,2-difluoro-ethanamine (1.2 g, 5.06 mmol, 1 eq.) and sodium bicarbonate (850 mg, 10.12 mmol, 2 eq.) in tetrahydrofuran (5 mL) and water (5 mL), di-tert-butyl dicarbonate (2.21 g, 10.12 mmol, 2.3 mL, 2 eq.) was added. The reaction mixture was stirred at 20 °C for 2 h. Water (100 mL) was added to the mixture, and the resulting mixture was extracted with ethyl acetate (50 mL x 3). The combined organic phase was washed with brine (60 mL), dried over sodium sulfate, filtered, and concentrated in vacuum. The residue was purified by silica gel chromatography (petroleum ether: ethyl acetate = 3:1). The crude product was purified by preparative high performance liquid chromatography (column: Phenomenex Gemini C18 250*50mm*10um; mobile phase: [water (0.05% ammonia hydroxide v/v)-ACN]; B%: 40%-65%, 22 min). tert-Butyl N-[1-(6-bromo-3-pyridyl)-2,2-difluoro-ethyl]carbamate (1.4g, 4.15mmol, 82% yield) was obtained as a pale yellow solid.
工程7
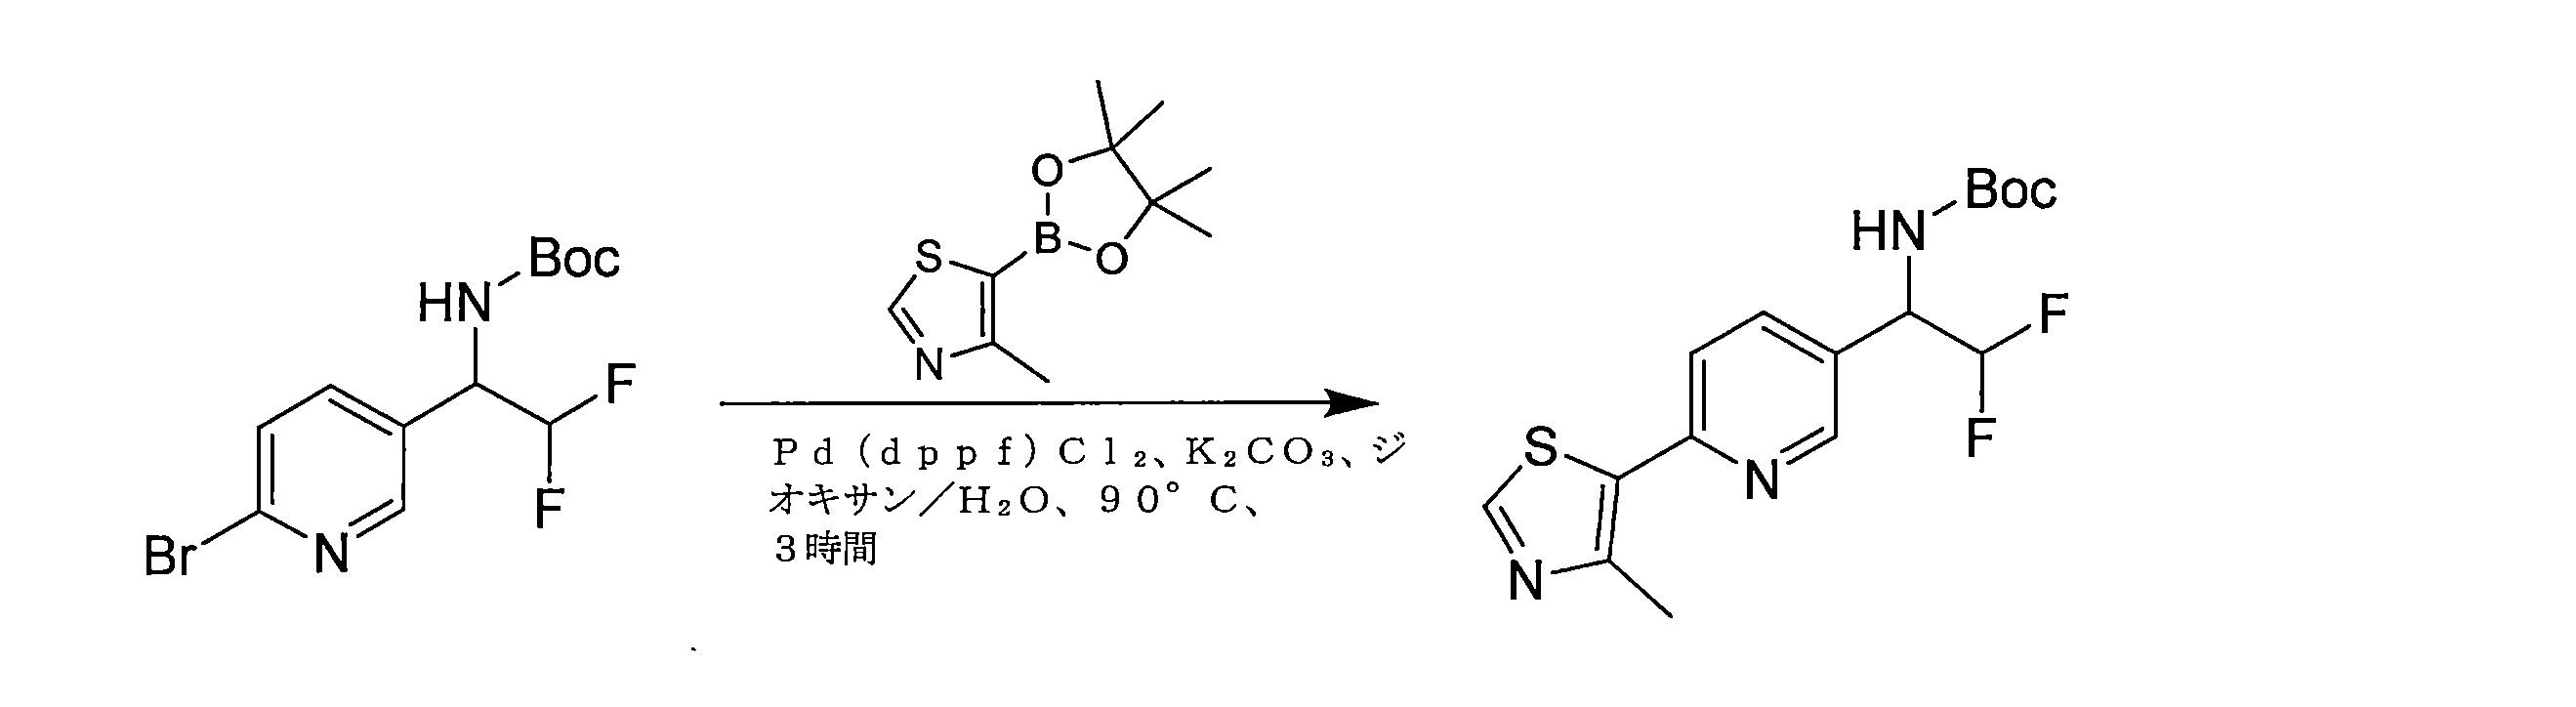
ジオキサン(50mL)および水(10mL)中のtert-ブチルN-[1-(6-ブロモ-3-ピリジル)-2,2-ジフルオロ-エチル]カルバメート(2g、5.93mmol、1当量)、4-メチル-5-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)チアゾール(3.34g、14.83mmol、2.5当量)の溶液に、炭酸カリウム(2.05g、14.83mmol、2.5当量)および(1,1’-ビス(ジフェニルホスフィノ)フェロセン)パラジウム(II)二塩化物(434mg、0.59mmol、0.1当量)を窒素下で添加した。反応混合物を100℃で12時間撹拌した。水(150mL)を混合物に添加し、得られた混合物を酢酸エチル(50mL×3)で抽出した。合わせた有機層をブライン(80mL)で洗浄し、硫酸ナトリウムで乾燥させ、濾過し、真空中で濃縮させた。残渣を、シリカゲルクロマトグラフィー(石油エーテル:酢酸エチル=100:1~1:1)によって精製した。次いで、残渣を、分取HPLC(カラム:Phenomenex Synergi C18 150*30mm*4um;移動相:[水(0.225%FA)-ACN];B%:35%~65%、10分)によってさらに精製した。tert-ブチルN-[2,2-ジフルオロ-1-[6-(4-メチルチアゾール-5-イル)-3-ピリジル]エチル]カルバメート(480mg、1.33mmol、収率22%)を淡い黄色の固体として得た。 To a solution of tert-butyl N-[1-(6-bromo-3-pyridyl)-2,2-difluoro-ethyl]carbamate (2 g, 5.93 mmol, 1 eq.), 4-methyl-5-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)thiazole (3.34 g, 14.83 mmol, 2.5 eq.) in dioxane (50 mL) and water (10 mL), potassium carbonate (2.05 g, 14.83 mmol, 2.5 eq.) and (1,1'-bis(diphenylphosphino)ferrocene)palladium(II) dichloride (434 mg, 0.59 mmol, 0.1 eq.) were added under nitrogen. The reaction mixture was stirred at 100 °C for 12 h. Water (150 mL) was added to the mixture and the resulting mixture was extracted with ethyl acetate (50 mL x 3). The combined organic layers were washed with brine (80 mL), dried over sodium sulfate, filtered, and concentrated in vacuo. The residue was purified by silica gel chromatography (petroleum ether:ethyl acetate=100:1-1:1). The residue was then further purified by preparative HPLC (column: Phenomenex Synergi C18 150*30 mm*4 um; mobile phase: [water (0.225% FA)-ACN]; B%: 35%-65%, 10 min). tert-Butyl N-[2,2-difluoro-1-[6-(4-methylthiazol-5-yl)-3-pyridyl]ethyl]carbamate (480 mg, 1.33 mmol, 22% yield) was obtained as a pale yellow solid.
工程8

tert-ブチルN-[2,2-ジフルオロ-1-[6-(4-メチルチアゾール-5-イル)-3-ピリジル]エチル]カルバメート(480mg、1.35mmol、1当量)を、分取SFC(SFCカラム:DAICEL CHIRALPAK AY 250×50mm、I.D.、10um;移動相:CO2中のエチルアルコール30%~30%;流速:60g/分;波長:220nm)によって精製した。異性体1(tert-ブチルN-[(1S)-2,2-ジフルオロ-1-[6-(4-メチルチアゾール-5-イル)-3-ピリジル]エチル]カルバメートとして暫定的に指定された;200mg、0.56mmol、収率82%)を淡い黄色の固体として得た。異性体2(tert-ブチルN-[(1R)-2,2-ジフルオロ-1-[6-(4-メチルチアゾール-5-イル)-3-ピリジル]エチル]カルバメートとして暫定的に指定された;210mg、0.59mmol、収率87%)を淡い黄色の固体として得た。 tert-Butyl N-[2,2-difluoro-1-[6-(4-methylthiazol-5-yl)-3-pyridyl]ethyl]carbamate (480 mg, 1.35 mmol, 1 equiv.) was purified by preparative SFC (SFC column: DAICEL CHIRALPAK AY 250×50 mm, I.D., 10 um; mobile phase: ethyl alcohol 30%-30% in CO2; flow rate: 60 g/min; wavelength: 220 nm). Isomer 1 (tentatively designated as tert-butyl N-[(1S)-2,2-difluoro-1-[6-(4-methylthiazol-5-yl)-3-pyridyl]ethyl]carbamate; 200 mg, 0.56 mmol, 82% yield) was obtained as a pale yellow solid. Isomer 2 (tentatively designated as tert-butyl N-[(1R)-2,2-difluoro-1-[6-(4-methylthiazol-5-yl)-3-pyridyl]ethyl]carbamate; 210 mg, 0.59 mmol, 87% yield) was obtained as a pale yellow solid.
工程9

N,N-ジメチルホルムアミド(5mL)中の2-[3-(2,2-ジエトキシエトキシ)イソキサゾール-5-イル]-3-メチル-ブタン酸(400mg、1.33mmol、1当量)およびメチル(2S,4R)-4-ヒドロキシピロリジン-2-カルボキシレート(241mg、1.33mmol、1当量、塩酸塩)の溶液に、N,N-ジイソプロピルエチルアミン(686mg、5.31mmol、0.9mL、4当量)およびO-(7-アザベンゾトリアゾール-1-イル)-N,N,N’,N’-テトラメチルウロニウムヘキサフルオロホスフェート(605mg、1.59mmol、1.2当量)を0℃で添加した。反応混合物を20℃で1時間撹拌した。水(50mL)を混合物に添加し、得られた混合物を酢酸エチル(20mL×3)で抽出した。合わせた有機相をブライン(30mL×2)で洗浄し、硫酸ナトリウムで乾燥させ、濾過し、真空中で濃縮した。残渣を、分取HPLC(カラム:Phenomenex Synergi C18 150*25*10um;移動相:[水(0.225%FA)-ACN];B%:36%~66%、10分)によって精製した。メチル(2S,4R)-1-[2-[3-(2,2-ジエトキシエトキシ)イソキサゾール-5-イル]-3-メチル-ブタノイル]-4-ヒドロキシ-ピロリジン-2-カルボキシレート(450mg、1.05mmol、収率78%)を無色の油として得た。 To a solution of 2-[3-(2,2-diethoxyethoxy)isoxazol-5-yl]-3-methyl-butanoic acid (400 mg, 1.33 mmol, 1 eq.) and methyl (2S,4R)-4-hydroxypyrrolidine-2-carboxylate (241 mg, 1.33 mmol, 1 eq., hydrochloride salt) in N,N-dimethylformamide (5 mL), N,N-diisopropylethylamine (686 mg, 5.31 mmol, 0.9 mL, 4 eq.) and O-(7-azabenzotriazol-1-yl)-N,N,N',N'-tetramethyluronium hexafluorophosphate (605 mg, 1.59 mmol, 1.2 eq.) were added at 0°C. The reaction mixture was stirred at 20°C for 1 h. Water (50 mL) was added to the mixture, and the resulting mixture was extracted with ethyl acetate (20 mL x 3). The combined organic phase was washed with brine (30 mL x 2), dried over sodium sulfate, filtered and concentrated in vacuum. The residue was purified by preparative HPLC (column: Phenomenex Synergi C18 150*25*10 um; mobile phase: [water (0.225% FA)-ACN]; B%: 36%-66%, 10 min). Methyl (2S,4R)-1-[2-[3-(2,2-diethoxyethoxy)isoxazol-5-yl]-3-methyl-butanoyl]-4-hydroxy-pyrrolidine-2-carboxylate (450 mg, 1.05 mmol, 78% yield) was obtained as a colorless oil.
工程10

混合物を、分取SFC(SFCカラム:DAICEL CHIRALPAK IC 250×30mm、I.D.、10um;移動相:CO2中のメタノール(0.1%NH4OH)30%~30%;流速:50g/分;波長:220nm)によって精製した。メチル(2S,4R)-1-[(2S)-2-[3-(2,2-ジエトキシエトキシ)イソキサゾール-5-イル]-3-メチル-ブタノイル]-4-ヒドロキシ-ピロリジン-2-カルボキシレート(180mg、0.36mmol、収率69%)を無色の油として得た。メチル(2S,4R)-1-[(2R)-2-[3-(2,2-ジエトキシエトキシ)イソキサゾール-5-イル]-3-メチル-ブタノイル]-4-ヒドロキシ-ピロリジン-2-カルボキシレート(220mg、0.51mmol、収率96%)を無色の油として得た。 The mixture was purified by preparative SFC (SFC column: DAICEL CHIRALPAK IC 250x30mm, I.D., 10um; mobile phase: methanol in CO2 (0.1% NH4OH) 30%-30%; flow rate: 50g/min; wavelength: 220nm). Methyl (2S,4R)-1-[(2S)-2-[3-(2,2-diethoxyethoxy)isoxazol-5-yl]-3-methyl-butanoyl]-4-hydroxy-pyrrolidine-2-carboxylate (180mg, 0.36mmol, 69% yield) was obtained as a colorless oil. Methyl (2S,4R)-1-[(2R)-2-[3-(2,2-diethoxyethoxy)isoxazol-5-yl]-3-methyl-butanoyl]-4-hydroxy-pyrrolidine-2-carboxylate (220 mg, 0.51 mmol, 96% yield) was obtained as a colorless oil.
他の実施例について上に記載されるものと類似の手順を使用して、以下のスキームに従って、メチル(2S,4R)-1-[(2R)-2-[3-(2,2-ジエトキシエトキシ)イソキサゾール-5-イル]-3-メチル-ブタノイル]-4-ヒドロキシ-ピロリジン-2-カルボキシレートを最終化合物に変換した。

例示的な化合物243を、類似の手順を使用して調製した。 Exemplary compound 243 was prepared using a similar procedure.
例示的な化合物246の例示的な合成
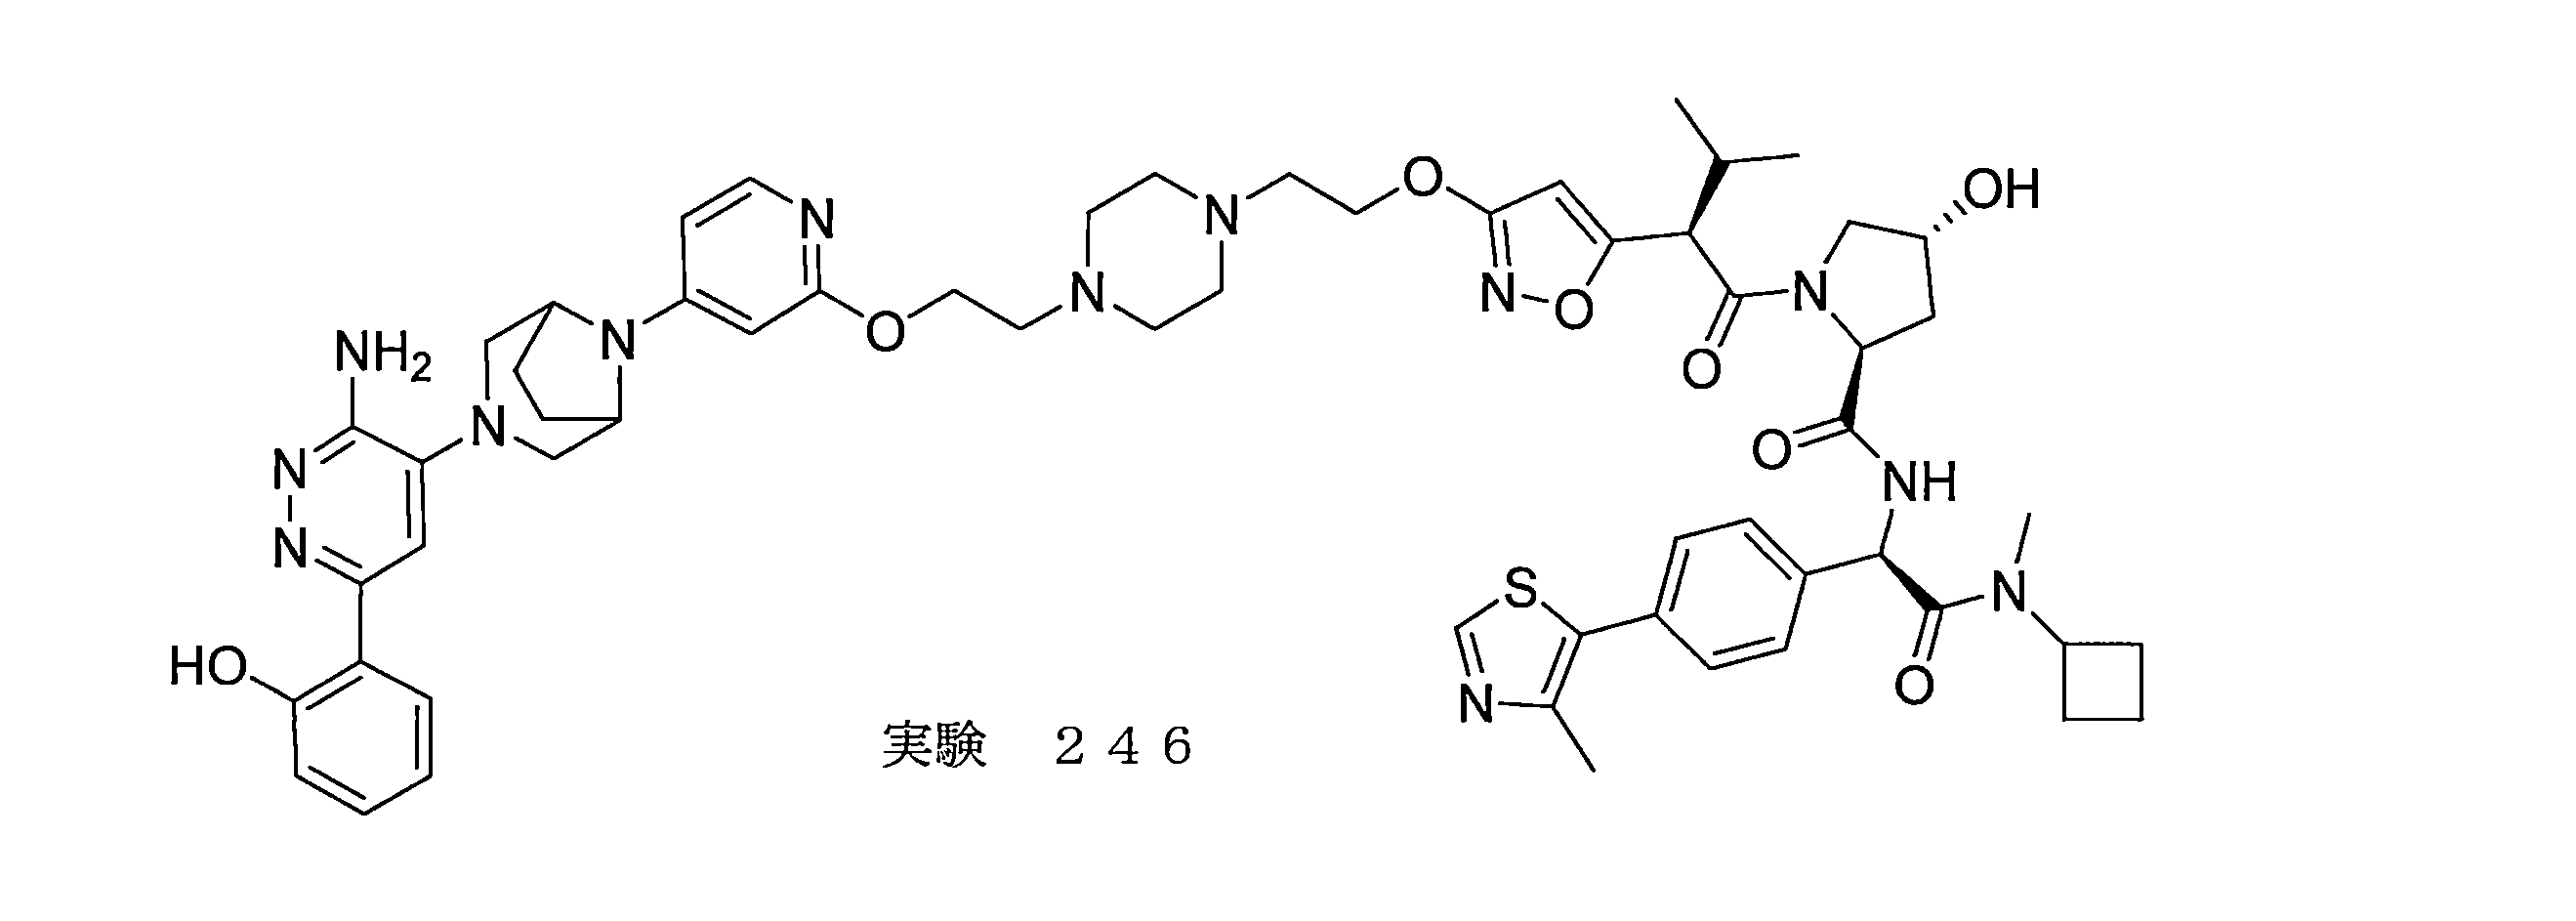
工程1

50mLの丸底フラスコに、DMF(15mL)中の(2R)-2-(4-ブロモフェニル)-2-[[(tert-ブトキシ)カルボニル]アミノ]酢酸(1g、3.029mmol、1当量)を入れ、それにジイソプロピルエチルアミン(1.57g、12.115mmol、4当量)、N-メチルシクロブタンアミン塩酸塩(368mg、3.03mmol、1当量)、およびBOP(1.61g、3.63mmol、1.2当量)を順番に添加した。得られた溶液を室温で2時間撹拌した。次いで、反応物を、水(10mL)を添加することによってクエンチした。得られた混合物を酢酸エチル(4×30mL)で抽出し、合わせた有機層をブライン(2×10mL)で洗浄し、無水硫酸ナトリウムで乾燥させ、濃縮した。残渣を、酢酸エチル/石油エーテル(1/1)で溶出するシリカゲルカラム上に適用した。これにより、1.1g(91%)のtert-ブチルN-[(R)-(4-ブロモフェニル)[シクロブチル(メチル)カルバモイル]メチル]カルバメートを黄色の固体として得た。 A 50 mL round bottom flask was charged with (2R)-2-(4-bromophenyl)-2-[[(tert-butoxy)carbonyl]amino]acetic acid (1 g, 3.029 mmol, 1 eq.) in DMF (15 mL), to which diisopropylethylamine (1.57 g, 12.115 mmol, 4 eq.), N-methylcyclobutanamine hydrochloride (368 mg, 3.03 mmol, 1 eq.), and BOP (1.61 g, 3.63 mmol, 1.2 eq.) were added in sequence. The resulting solution was stirred at room temperature for 2 h. The reaction was then quenched by adding water (10 mL). The resulting mixture was extracted with ethyl acetate (4 x 30 mL) and the combined organic layers were washed with brine (2 x 10 mL), dried over anhydrous sodium sulfate, and concentrated. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (1/1). This yielded 1.1 g (91%) of tert-butyl N-[(R)-(4-bromophenyl)[cyclobutyl(methyl)carbamoyl]methyl]carbamate as a yellow solid.
工程2

100mLの丸底フラスコに、ジオキサン(10mL)およびH2O(2mL)中のtert-ブチルN-[(R)-(4-ブロモフェニル)[シクロブチル(メチル)カルバモイル]メチル]カルバメート(1g、2.52mmol、1当量)、K2CO3(1.04g、7.55mmol、3当量)、4-メチル-5-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)-1,3-チアゾール(850mg、3.78mmol、1.5当量)、およびPd(dppf)Cl2(184mg、0.25mmol、0.1当量)を窒素雰囲気下で入れた。得られた混合物を、油浴中で、窒素雰囲気下で、90℃で3時間撹拌した。混合物を濃縮し、残渣を、酢酸エチル/石油エーテル(1/1)で溶出するシリカゲルカラム上に適用した。これにより、480mg(46%)のtert-ブチルN-[(R)-[シクロブチル(メチル)カルバモイル][4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]カルバメートを黄色の固体として得た。 A 100 mL round-bottom flask was charged with tert-butyl N-[(R)-(4-bromophenyl)[cyclobutyl(methyl)carbamoyl]methyl]carbamate (1 g, 2.52 mmol, 1 eq.), K2CO3 (1.04 g, 7.55 mmol, 3 eq.), 4-methyl-5-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)-1,3-thiazole (850 mg, 3.78 mmol, 1.5 eq.), and Pd(dppf)Cl2 (184 mg, 0.25 mmol, 0.1 eq.) in dioxane (10 mL) and H2O (2 mL) under nitrogen atmosphere. The resulting mixture was stirred at 90 °C in an oil bath under nitrogen atmosphere for 3 h. The mixture was concentrated and the residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (1/1). This yielded 480 mg (46%) of tert-butyl N-[(R)-[cyclobutyl(methyl)carbamoyl][4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]carbamate as a yellow solid.
他の実施例について上に記載されるものと類似の手順を使用して、tert-ブチルN-[(R)-[シクロブチル(メチル)カルバモイル][4-(4-メチル-1,3-チアゾール-5-イル)フェニル]メチル]カルバメートを最終化合物に変換した。 Using procedures similar to those described above for other examples, tert-butyl N-[(R)-[cyclobutyl(methyl)carbamoyl][4-(4-methyl-1,3-thiazol-5-yl)phenyl]methyl]carbamate was converted to the final compound.
例示的な化合物245を、類似の手順を使用して調製した。 Exemplary compound 245 was prepared using a similar procedure.
例示的な化合物248の例示的な合成
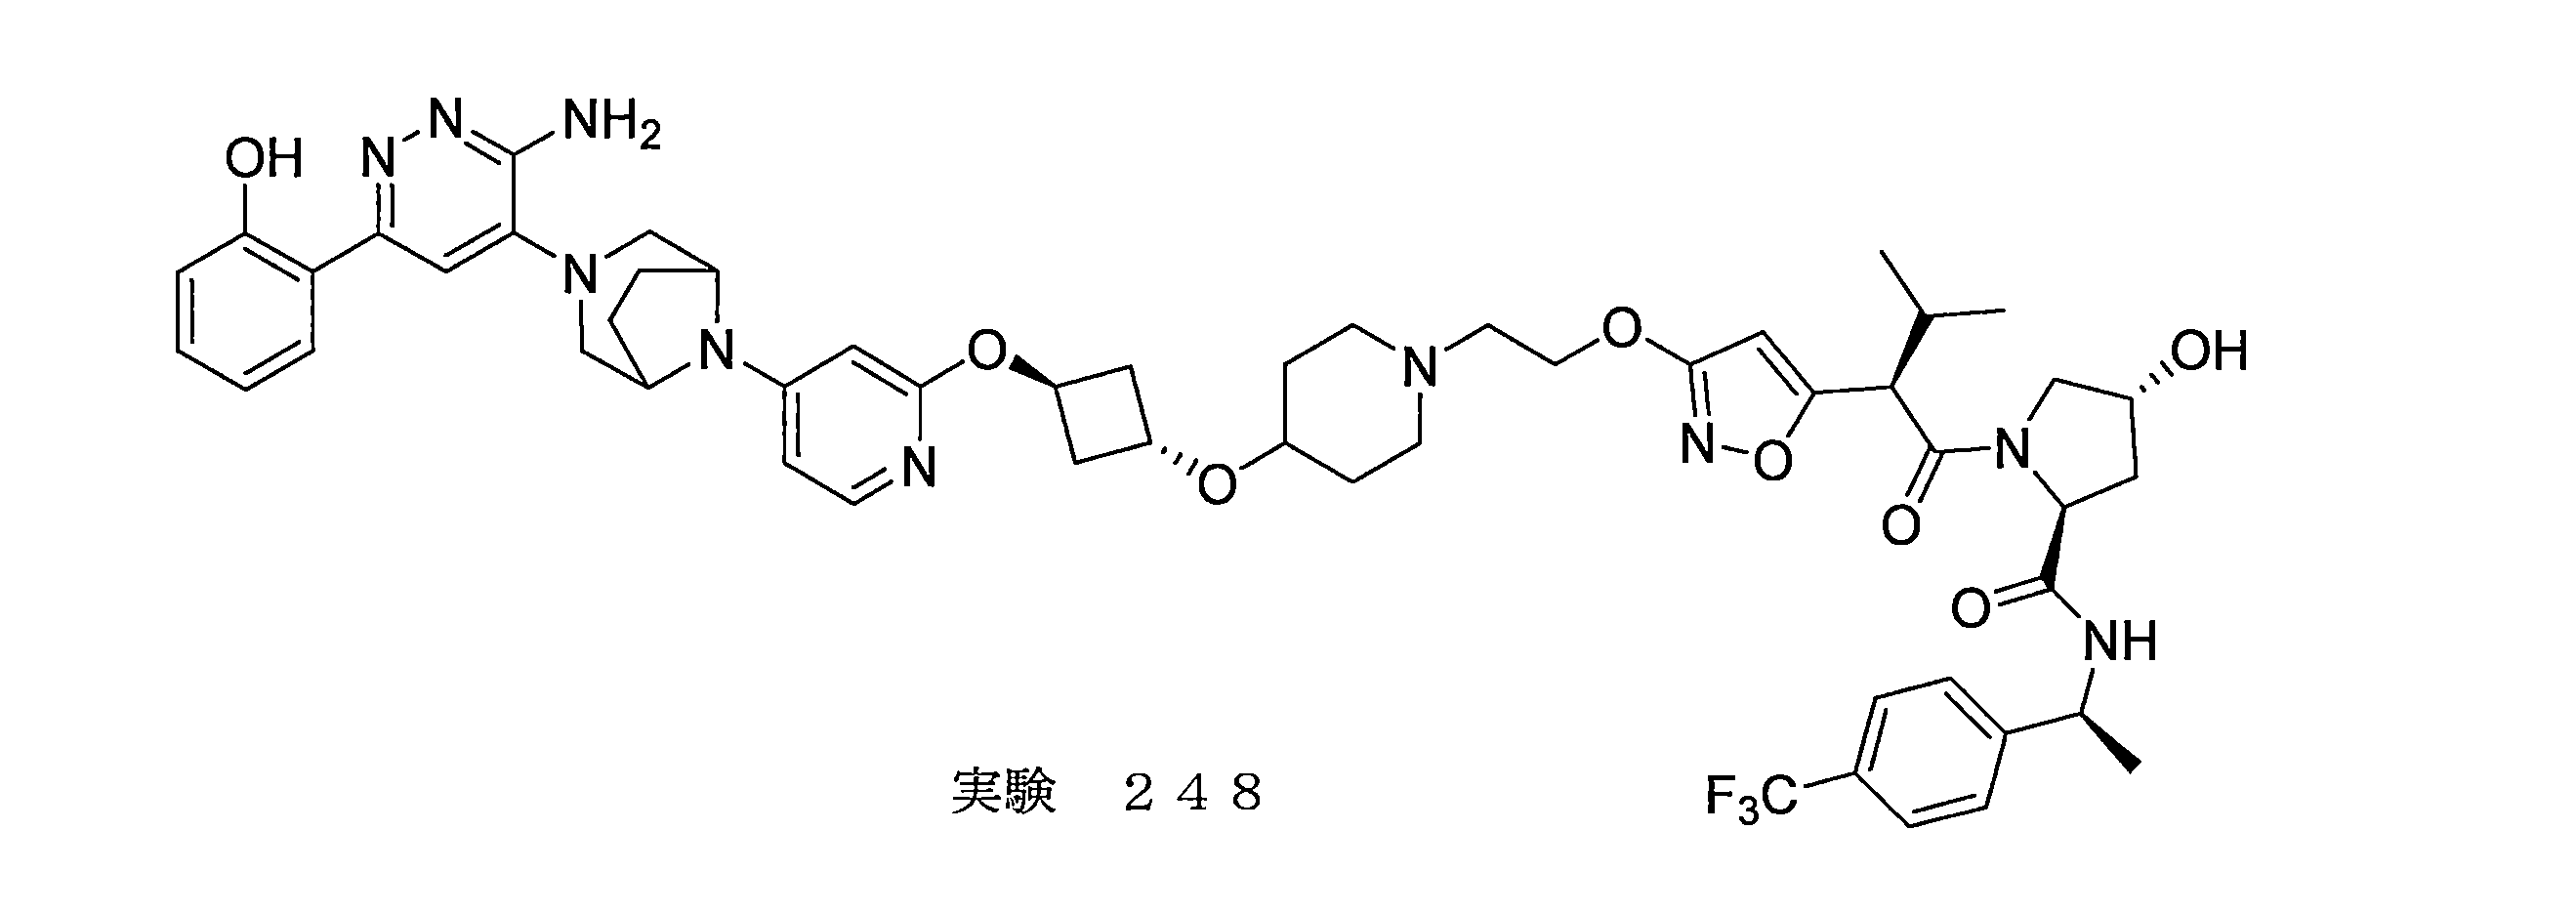
工程1

250のmL丸底フラスコに、THF(50mL)中の4-(トリフルオロメチル)ベンズアルデヒド(3.0g、17.2mmol、1.0当量)、(S)-2-メチルプロパン-2-スルフィンアミド(4.2g、0.035mmol、2.0当量)、およびチタンイソプロポキシド(14.9g、0.052mmol、3.0当量)の溶液を入れた。得られた溶液、油浴中で、80℃で16時間撹拌した。次いで、反応物を、200mLの水を添加することによってクエンチし、混合物を酢酸エチル(100mL×3)で抽出した。合わせた有機相をブラインで洗浄し、硫酸ナトリウムで乾燥させ、減圧下で濃縮した。残渣を、酢酸エチル/石油エーテル(1:1)で溶出するシリカゲルカラム上に適用した。これにより、3.1g(65%)の(S)-2-メチル-N-[[4-(トリフルオロメチル)フェニル]メチリデン]プロパン-2-スルフィンアミドを黄色の油として得た。 A 250 mL round-bottom flask was charged with a solution of 4-(trifluoromethyl)benzaldehyde (3.0 g, 17.2 mmol, 1.0 equiv.), (S)-2-methylpropane-2-sulfinamide (4.2 g, 0.035 mmol, 2.0 equiv.), and titanium isopropoxide (14.9 g, 0.052 mmol, 3.0 equiv.) in THF (50 mL). The resulting solution was stirred at 80 °C in an oil bath for 16 h. The reaction was then quenched by adding 200 mL of water, and the mixture was extracted with ethyl acetate (100 mL x 3). The combined organic phase was washed with brine, dried over sodium sulfate, and concentrated under reduced pressure. The residue was applied onto a silica gel column eluting with ethyl acetate/petroleum ether (1:1). This yielded 3.1 g (65%) of (S)-2-methyl-N-[[4-(trifluoromethyl)phenyl]methylidene]propane-2-sulfinamide as a yellow oil.
工程2
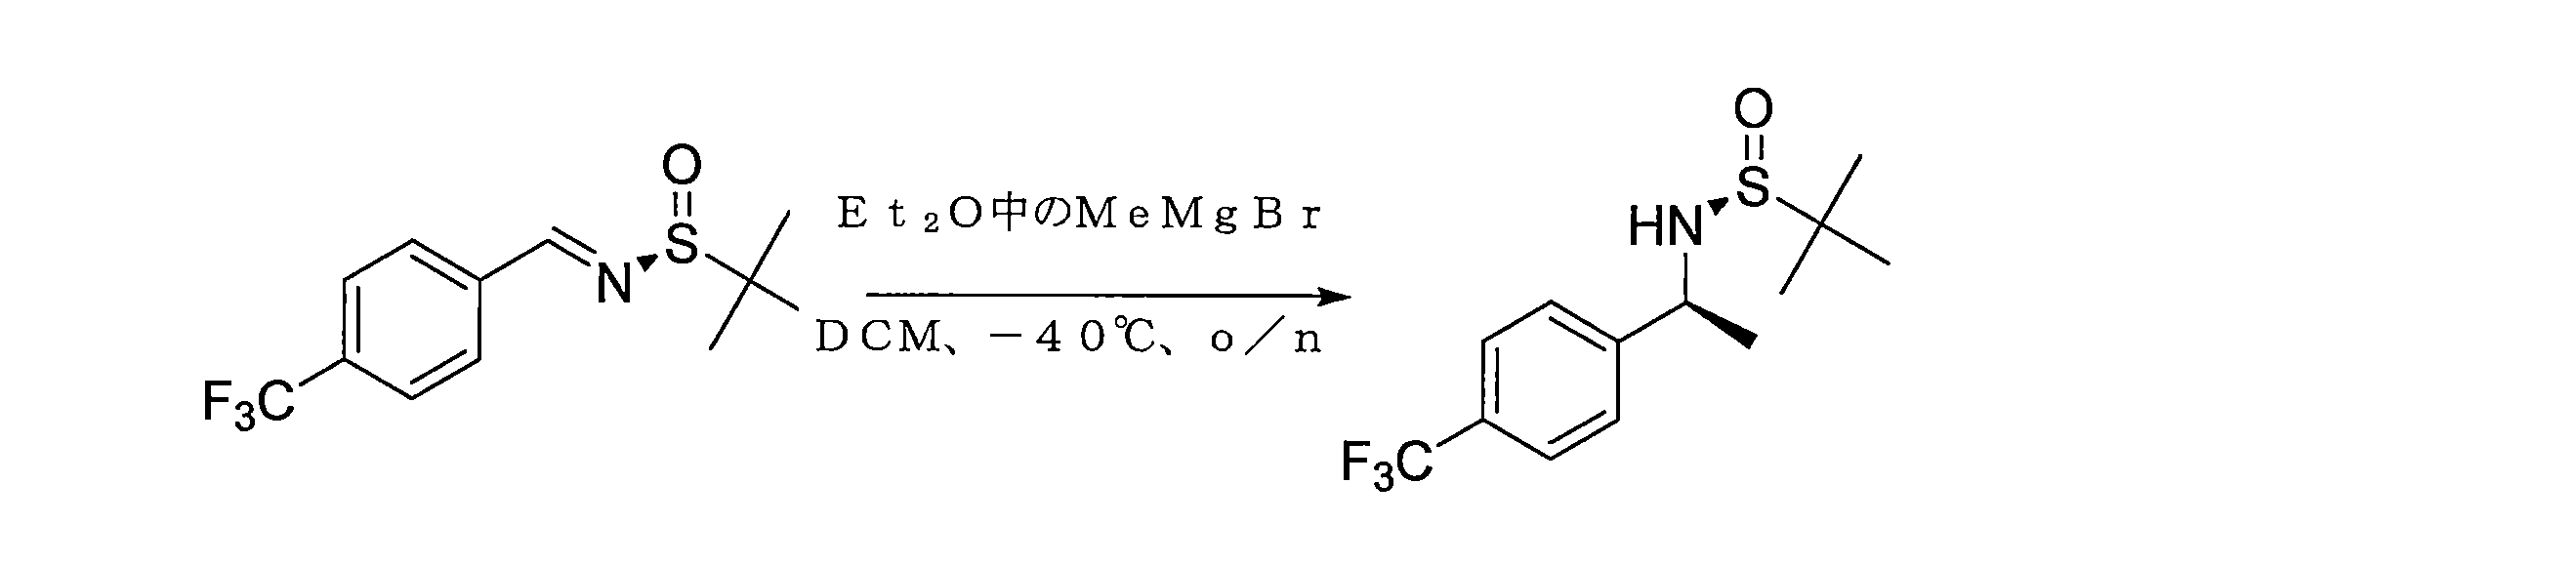
DCM(20mL)中の(S)-2-メチル-N-[[4-(トリフルオロメチル)フェニル]メチリデン]プロパン-2-スルフィンアミド(3.1g、11.2mmol、1.0当量)の溶液に、メチルマグネシウムクロリド溶液(エーテル中3.0M、5mL)を窒素雰囲気下で、-48℃で20分の期間にわたって滴下で添加した。次いで、得られた混合物を室温で2時間撹拌した。次いで、反応混合物を、50mLの飽和NH4Cl溶液を添加することによってクエンチし、ジクロロメタン(50mL×3)で抽出した。合わせた有機相をブラインで洗浄し、硫酸ナトリウムで乾燥させ、減圧下で濃縮した。残渣を、酢酸エチル/石油エーテル(1:1)で溶出するシリカゲルカラム上に適用した。これにより、1.83g(56%)の(S)-2-メチル-N-[(1S)-1-[4-(トリフルオロメチル)フェニル]エチル]プロパン-2-スルフィンアミドを白色の固体として得た。 To a solution of (S)-2-methyl-N-[[4-(trifluoromethyl)phenyl]methylidene]propane-2-sulfinamide (3.1 g, 11.2 mmol, 1.0 equiv.) in DCM (20 mL) was added dropwise over a period of 20 min at −48° C. under nitrogen atmosphere with methylmagnesium chloride solution (3.0 M in ether, 5 mL). The resulting mixture was then stirred at room temperature for 2 h. The reaction mixture was then quenched by adding 50 mL of saturated NH4Cl solution and extracted with dichloromethane (50 mL×3). The combined organic phase was washed with brine, dried over sodium sulfate and concentrated under reduced pressure. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (1:1). This gave 1.83 g (56%) of (S)-2-methyl-N-[(1S)-1-[4-(trifluoromethyl)phenyl]ethyl]propane-2-sulfinamide as a white solid.
工程3

100mLの丸底フラスコに、メタノール(10mL)中の(S)-2-メチル-N-[(1S)-1-[4-(トリフルオロメチル)フェニル]エチル]プロパン-2-スルフィンアミド(1.8g、6.2mmol、1.0当量)を入れた。続いて、塩化水素水溶液(37%、5mL)を添加した。得られた溶液を室温で2時間撹拌し、次いで、減圧下で濃縮した。これにより、970mg(82%)の(1S)-1-[4-(トリフルオロメチル)フェニル]エタン-1-アミンを白色の固体として得た。 A 100 mL round-bottom flask was charged with (S)-2-methyl-N-[(1S)-1-[4-(trifluoromethyl)phenyl]ethyl]propane-2-sulfinamide (1.8 g, 6.2 mmol, 1.0 equiv.) in methanol (10 mL). Aqueous hydrogen chloride (37%, 5 mL) was then added. The resulting solution was stirred at room temperature for 2 h and then concentrated under reduced pressure. This afforded 970 mg (82%) of (1S)-1-[4-(trifluoromethyl)phenyl]ethan-1-amine as a white solid.
他の実施例について上に記載されるものと類似の手順を使用して、(1S)-1-[4-(トリフルオロメチル)フェニル]エタン-1-アミンを最終化合物に変換した。 (1S)-1-[4-(trifluoromethyl)phenyl]ethan-1-amine was converted to the final compound using procedures similar to those described above for other examples.
実施例の化合物247を、類似の手順を使用して調製した。 Example compound 247 was prepared using a similar procedure.
例示的な化合物404の例示的な合成
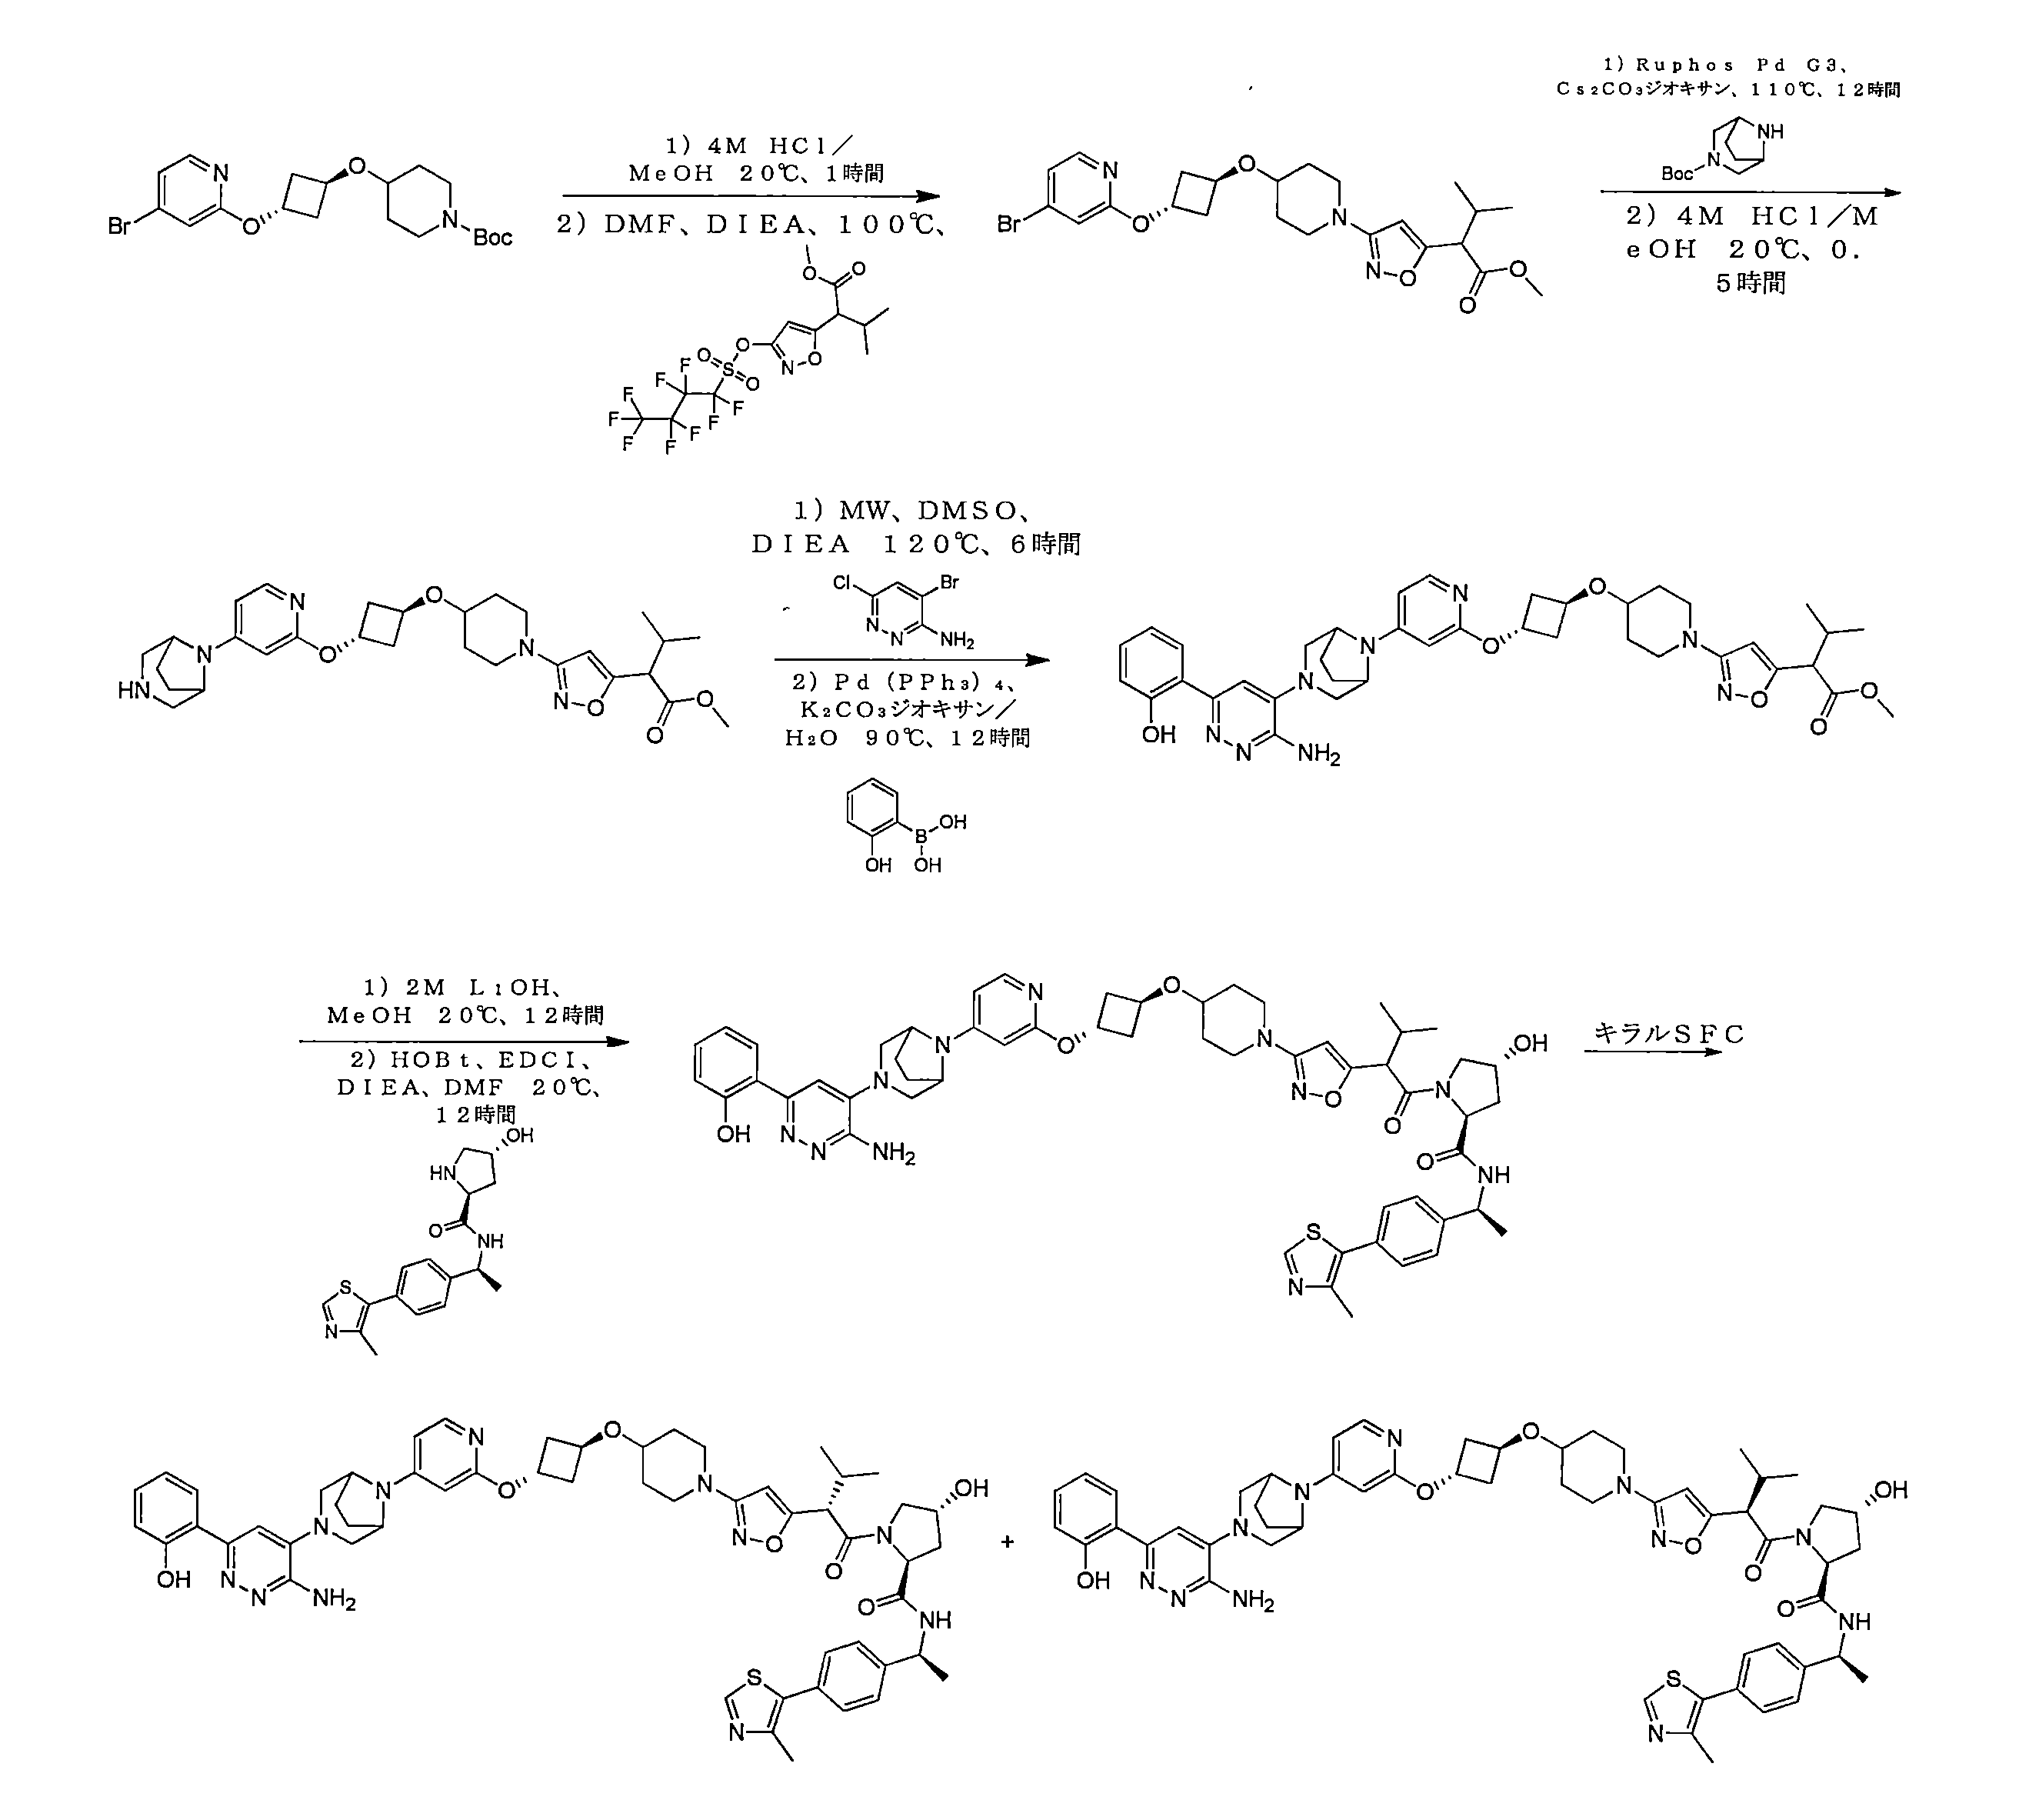
上記のスキームに示されるように、tert-ブチル4-((1R,3R)-3-((4-ブロモピリジン-2-イル)オキシ)シクロブトキシ)ピペリジン-1-カルボキシレートを、酸性条件下で脱保護し、得られた遊離環状アミンを活性化イソキサゾール誘導体と反応させて、メチル3-メチル-2-(3-(((ペルフルオロブチル)スルホニル)オキシ)イソキサゾール-5-イル)ブタノエートをカップリング生成物として形成した。tert-ブチル3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレートとの、生成物中の臭化ピリジル断片のパラジウム触媒化反応、続いてtert-ブチルオキシカルボニル(Boc)保護基の切断は、遊離アミンを産出し、それはアミンの存在下で、DMSO中の加熱下で、4-ブロモ-6-クロロピリダジン-3-アミンとの求核カップリング反応を受けた。ピリダジン臭化物と(2-ヒドロキシフェニル)ボロン酸とのスズキカップリング反応は、PTM結合部分のアセンブリを完了した。エステル加水分解、続いて(2S,4R)-4-ヒドロキシ-N-((S)-1-(4-(4-メチルチアゾール-5-イル)フェニル)エチル)ピロリジン-2-カルボキサミドとの縮合により、ジアステレオマーの混合物として所望の最終生成物を得、それを超臨界流体クロマトグラフィー(SFC)によって分離した。 As shown in the scheme above, tert-butyl 4-((1R,3R)-3-((4-bromopyridin-2-yl)oxy)cyclobutoxy)piperidine-1-carboxylate was deprotected under acidic conditions and the resulting free cyclic amine was reacted with an activated isoxazole derivative to form methyl 3-methyl-2-(3-(((perfluorobutyl)sulfonyl)oxy)isoxazol-5-yl)butanoate as the coupling product. Palladium-catalyzed reaction of the pyridyl bromide fragment in the product with tert-butyl 3,8-diazabicyclo[3.2.1]octane-3-carboxylate, followed by cleavage of the tert-butyloxycarbonyl (Boc) protecting group, yielded the free amine, which underwent nucleophilic coupling reaction with 4-bromo-6-chloropyridazin-3-amine under heating in DMSO in the presence of an amine. Suzuki coupling reaction of pyridazine bromide with (2-hydroxyphenyl)boronic acid completed the assembly of the PTM binding moiety. Ester hydrolysis followed by condensation with (2S,4R)-4-hydroxy-N-((S)-1-(4-(4-methylthiazol-5-yl)phenyl)ethyl)pyrrolidine-2-carboxamide gave the desired final products as a mixture of diastereomers, which were separated by supercritical fluid chromatography (SFC).
例示的な化合物411の例示的な合成
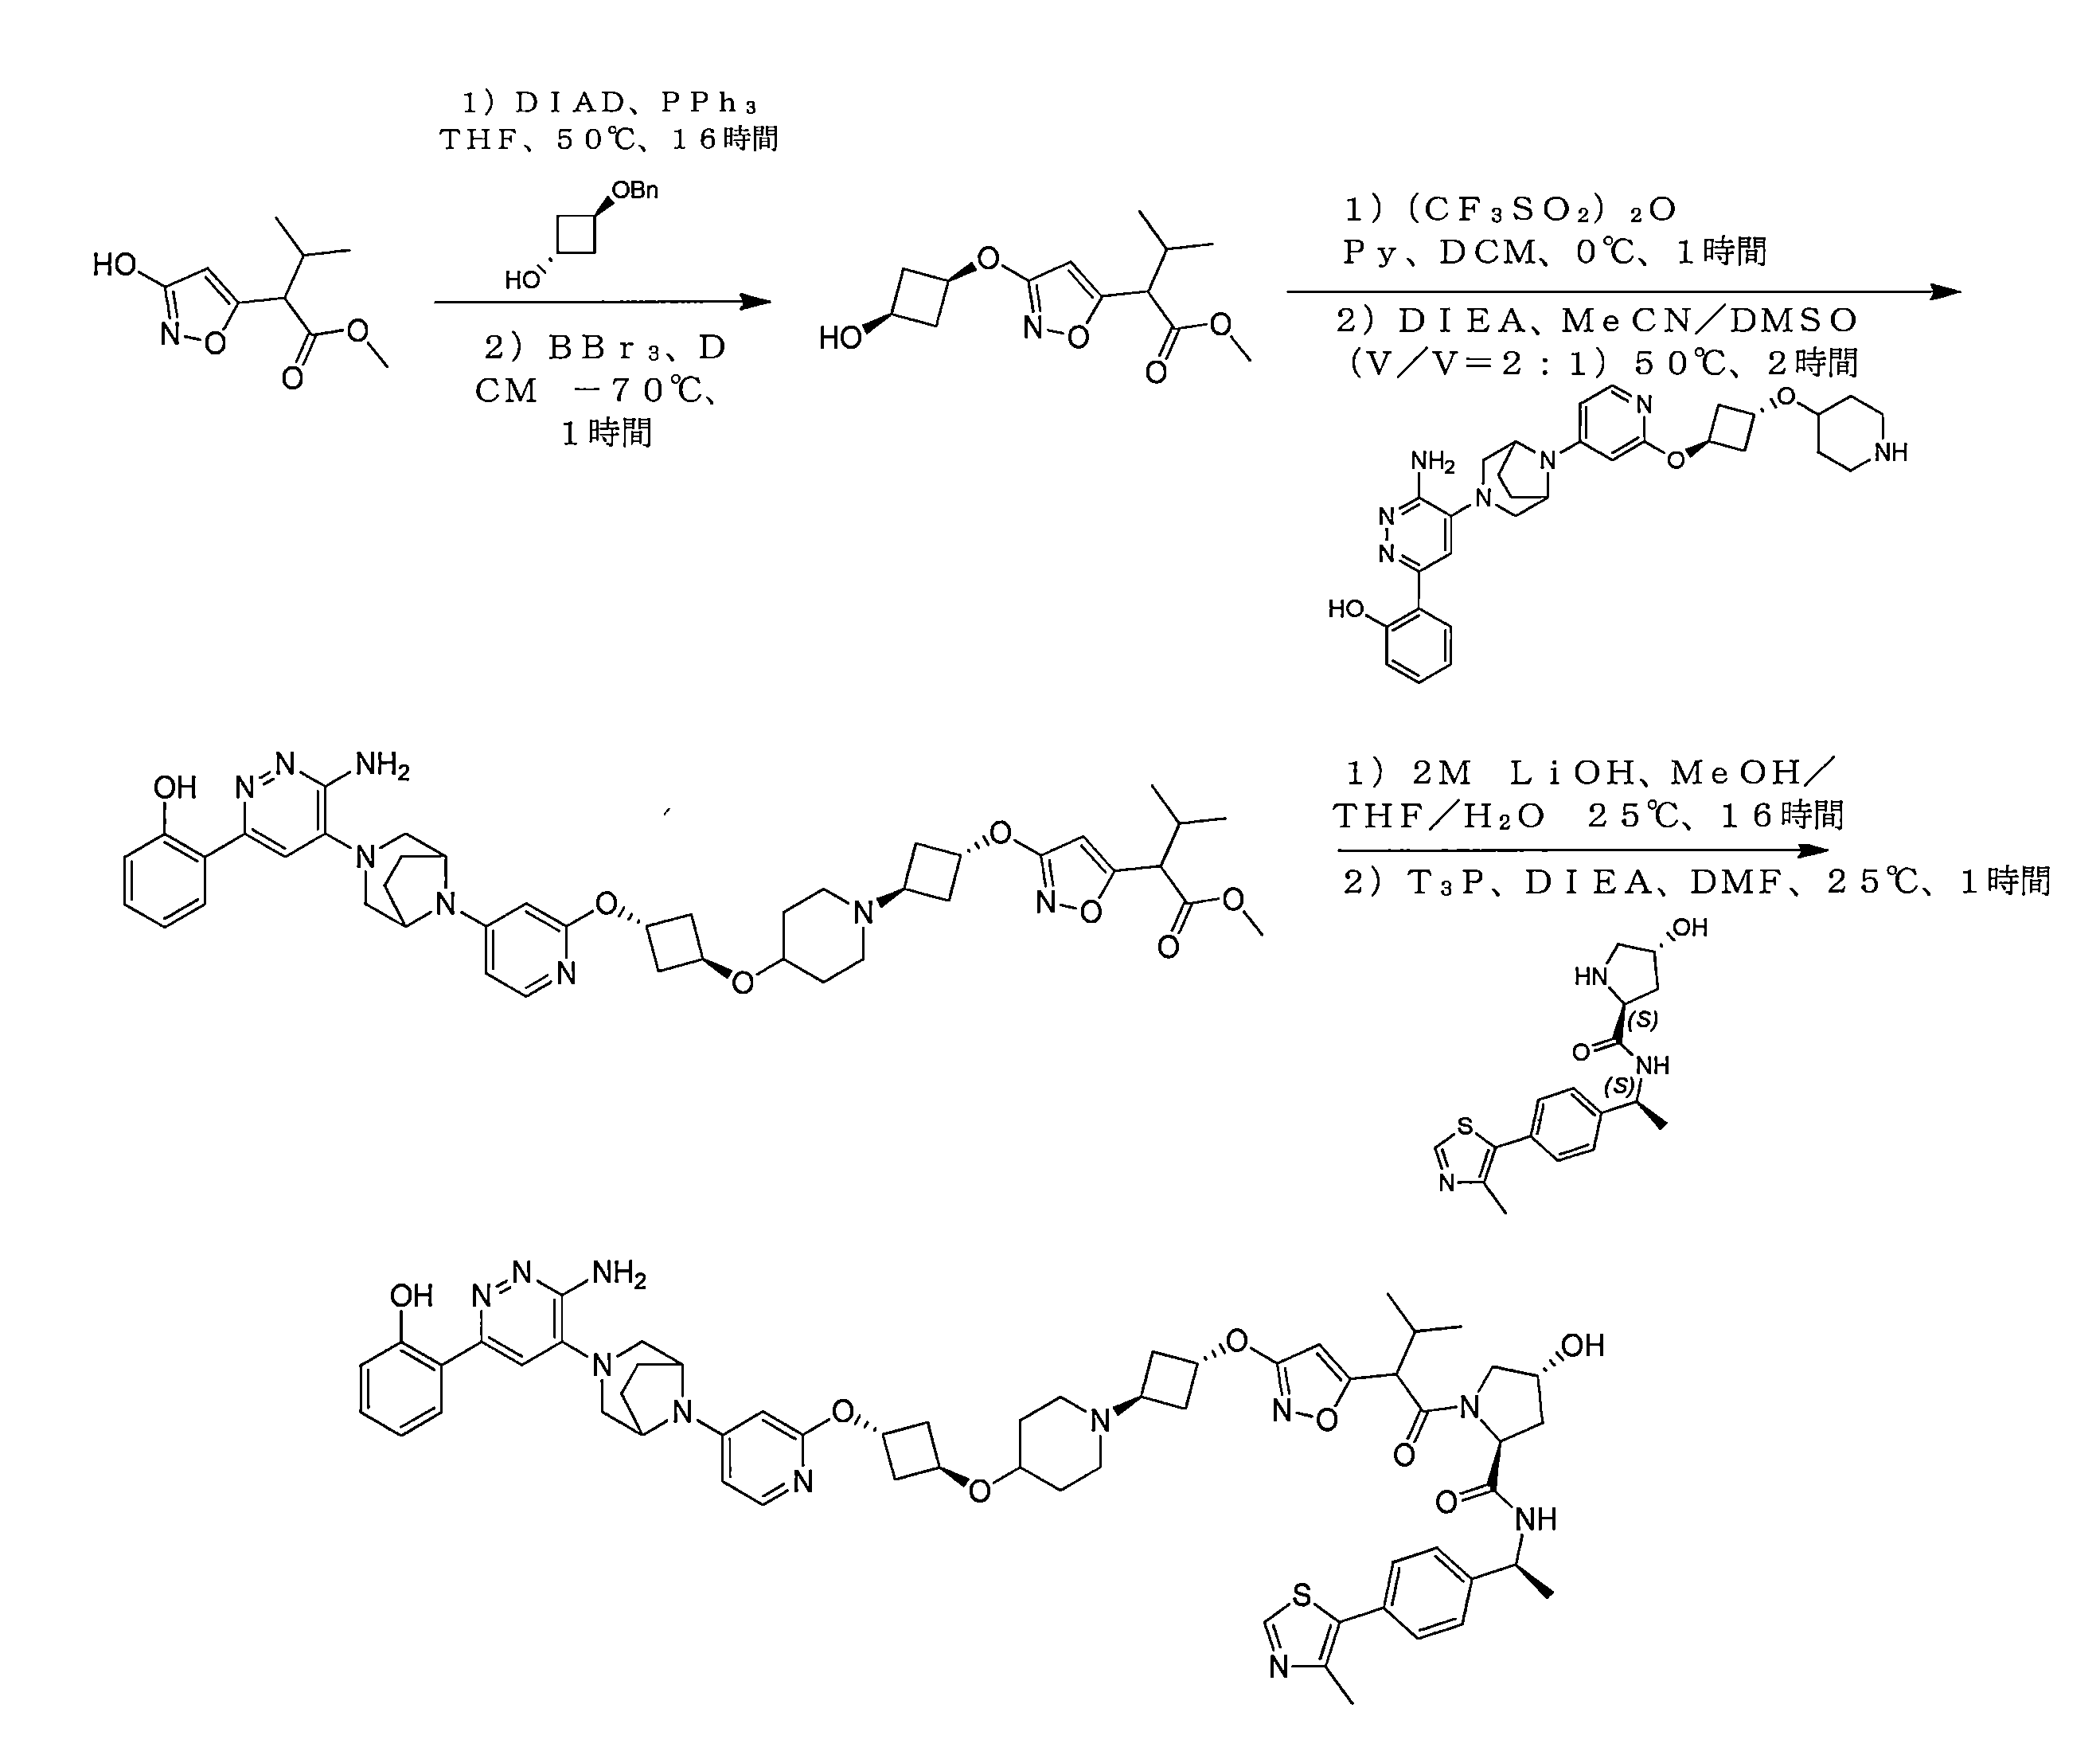
上記のスキームに示されるように、メチル2-(3-ヒドロキシイソキサゾール-5-イル)-3-メチルブタノエートおよび(1R,3R)-3-(ベンジルオキシ)シクロブタノールの間のミツノブ反応、続いてベンジルオキシ基切断は、メチル2-(3-((1S,3S)-3-ヒドロキシシクロブトキシ)イソキサゾール-5-イル)-3-メチルブタノエートを提供した。トリフルオロメタンスルホン酸無水物でのヒドロキシ基の活性化、続くWPTM5-WPTM3-WPTM2-WPTM1断片を含む環状アミンでのトリフラート置換により、結合基Lを含んだカップリング生成物を得た。エステル加水分解、続く(2S,4R)-4-ヒドロキシ-N-((S)-1-(4-(4-メチルチアゾール-5-イル)フェニル)エチル)ピロリジン-2-カルボキサミドでの縮合により、所望の最終生成物を得た。 As shown in the scheme above, Mitsunobu reaction between methyl 2-(3-hydroxyisoxazol-5-yl)-3-methylbutanoate and (1R,3R)-3-(benzyloxy)cyclobutanol, followed by benzyloxy group cleavage, provided methyl 2-(3-((1S,3S)-3-hydroxycyclobutoxy)isoxazol-5-yl)-3-methylbutanoate. Activation of the hydroxy group with trifluoromethanesulfonic anhydride, followed by triflate displacement with a cyclic amine containing WPTM5-WPTM3-WPTM2-WPTM1 fragment, afforded the coupling product containing linking group L. Ester hydrolysis, followed by condensation with (2S,4R)-4-hydroxy-N-((S)-1-(4-(4-methylthiazol-5-yl)phenyl)ethyl)pyrrolidine-2-carboxamide afforded the desired final product.
例示的な化合物428の例示的な合成

上記のスキームに示されるように、4-ブロモ-6-クロロピリダジン-3-アミンを4-ビニル誘導体に変換し、それはtert-ブチル4-((1R,3R)-3-((4-ブロモピリジン-2-イル)オキシ)シクロブトキシ)ピペリジン-1-カルボキシレートとのヘックカップリング反応を受けて、カップリング生成物を生成した。次いで、生成物中の6-クロロピリダジン部分は、スズキ条件下で(2-ヒドロキシフェニル)ボロン酸と反応して、WPTM5-WPTM3断片をアセンブリした。その後のBoc脱保護、続いて(2S,4R)-4-ヒドロキシ-1-((R)-3-メチル-2-(3-(2-オキソエトキシ)イソキサゾール-5-イル)ブタノイル)-N-((S)-1-(4-(4-メチルチアゾール-5-イル)フェニル)エチル)ピロリジン-2-カルボキサミドによる還元的アミノ化は、所望の最終生成物を提供した。 As shown in the scheme above, 4-bromo-6-chloropyridazin-3-amine was converted to the 4-vinyl derivative, which underwent a Heck coupling reaction with tert-butyl 4-((1R,3R)-3-((4-bromopyridin-2-yl)oxy)cyclobutoxy)piperidine-1-carboxylate to generate the coupling product. The 6-chloropyridazine moiety in the product was then reacted with (2-hydroxyphenyl)boronic acid under Suzuki conditions to assemble the WPTM5-WPTM3 fragment. Subsequent Boc deprotection followed by reductive amination with (2S,4R)-4-hydroxy-1-((R)-3-methyl-2-(3-(2-oxoethoxy)isoxazol-5-yl)butanoyl)-N-((S)-1-(4-(4-methylthiazol-5-yl)phenyl)ethyl)pyrrolidine-2-carboxamide provided the desired final product.
例示的な化合物445の例示的な合成
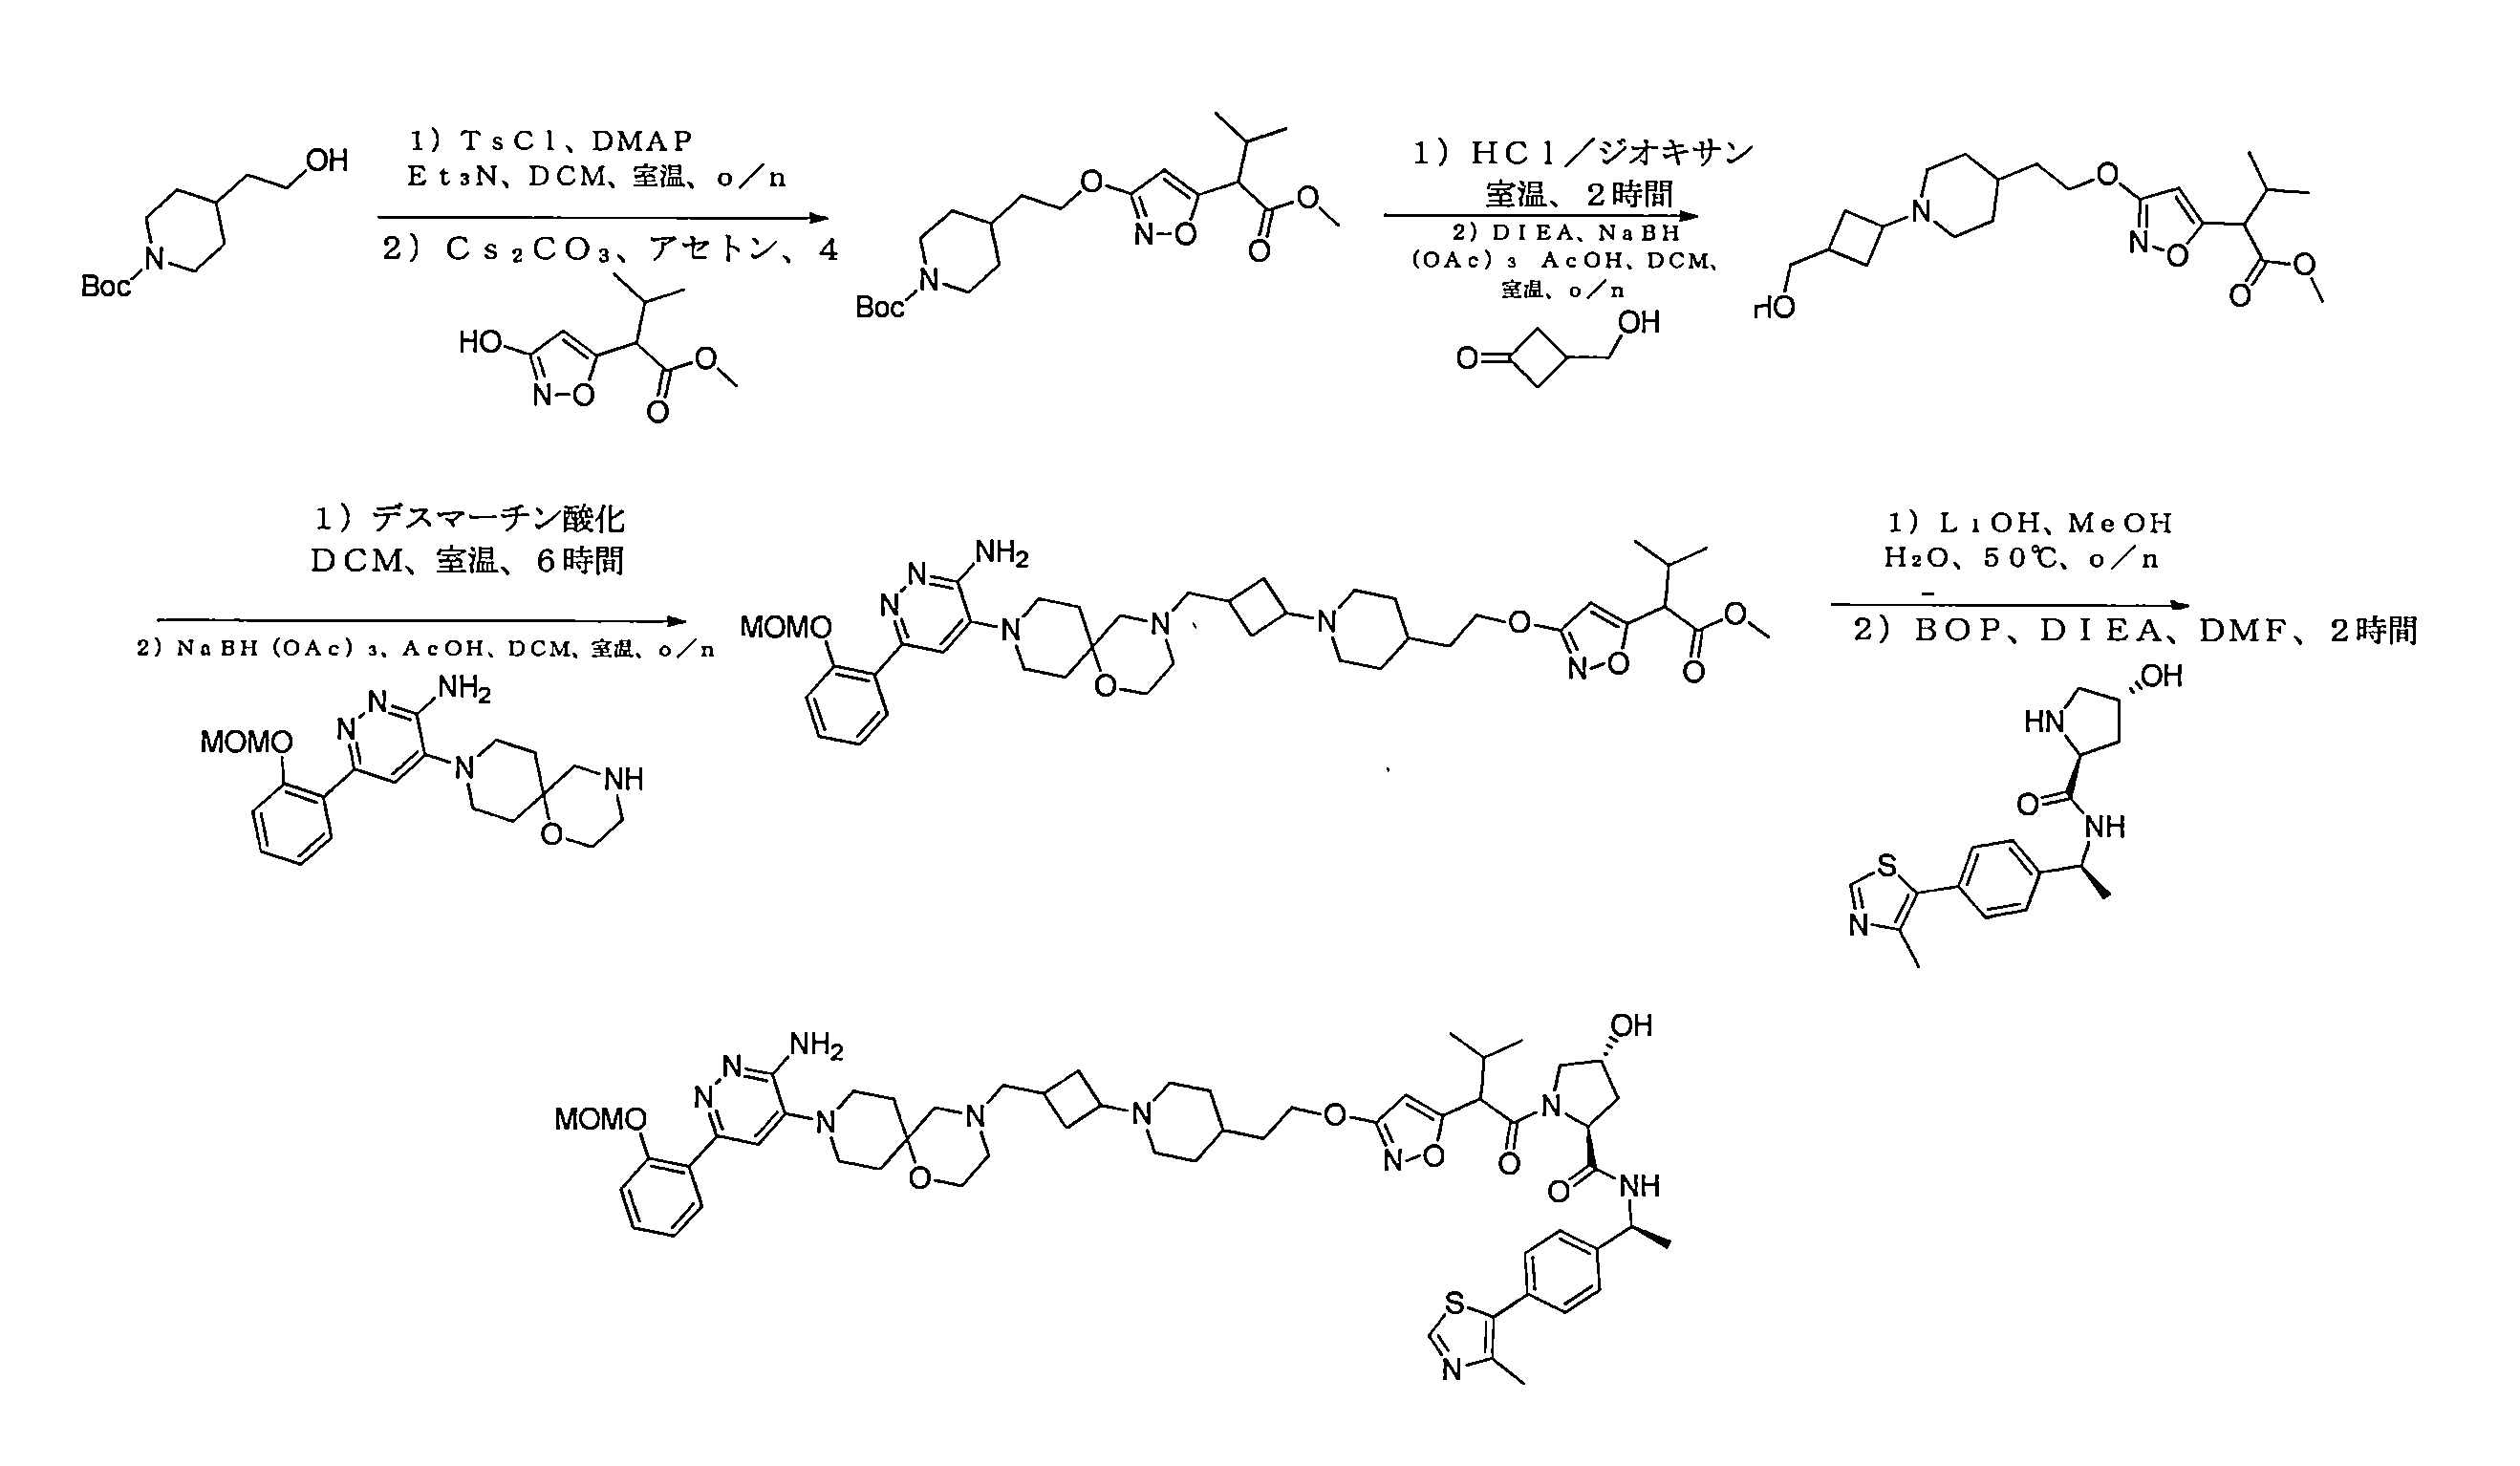
上記のスキームに示されるように、tert-ブチル4-(2-ヒドロキシエチル)ピペリジン-1-カルボキシレート中のヒドロキシル基を塩化トシルで活性化し、得られたトシレートを2-(3-ヒドロキシイソキサゾール-5-イル)-3-メチルブタン酸メチルで求核置換した。生成物中のBoc基を切断し、遊離環状アミンを3-(ヒドロキシメチル)シクロブタノンおよびトリアセトキシ水素化ホウ素ナトリウムと反応させて、2-(3-(2-(1-(3-(ヒドロキシメチル)シクロブチル)ピペリジン-4-イル)エトキシ)イソキサゾール-5-イル)-3-メチルブタン酸メチルを形成した。ヒドロキシル基のデスマーチン酸化、続くスピロ-アミンでの得られたアルデヒドの還元的アミノ化により、PTM結合部分および連結基Lを組み合わせたカップリング生成物を得た。エステル加水分解、続く(2S,4R)-4-ヒドロキシ-N-((S)-1-(4-(4-メチルチアゾール-5-イル)フェニル)エチル)ピロリジン-2-カルボキサミドでの縮合により、所望の最終生成物を得た。 As shown in the scheme above, the hydroxyl group in tert-butyl 4-(2-hydroxyethyl)piperidine-1-carboxylate was activated with tosyl chloride and the resulting tosylate was nucleophilically substituted with methyl 2-(3-hydroxyisoxazol-5-yl)-3-methylbutanoate. The Boc group in the product was cleaved and the free cyclic amine was reacted with 3-(hydroxymethyl)cyclobutanone and sodium triacetoxyborohydride to form methyl 2-(3-(2-(1-(3-(hydroxymethyl)cyclobutyl)piperidin-4-yl)ethoxy)isoxazol-5-yl)-3-methylbutanoate. Dess-Martin oxidation of the hydroxyl group followed by reductive amination of the resulting aldehyde with a spiro-amine afforded the coupled product combining the PTM binding moiety and the linking group L. Ester hydrolysis followed by condensation with (2S,4R)-4-hydroxy-N-((S)-1-(4-(4-methylthiazol-5-yl)phenyl)ethyl)pyrrolidine-2-carboxamide gave the desired final product.
例示的な化合物449~456の例示的な合成
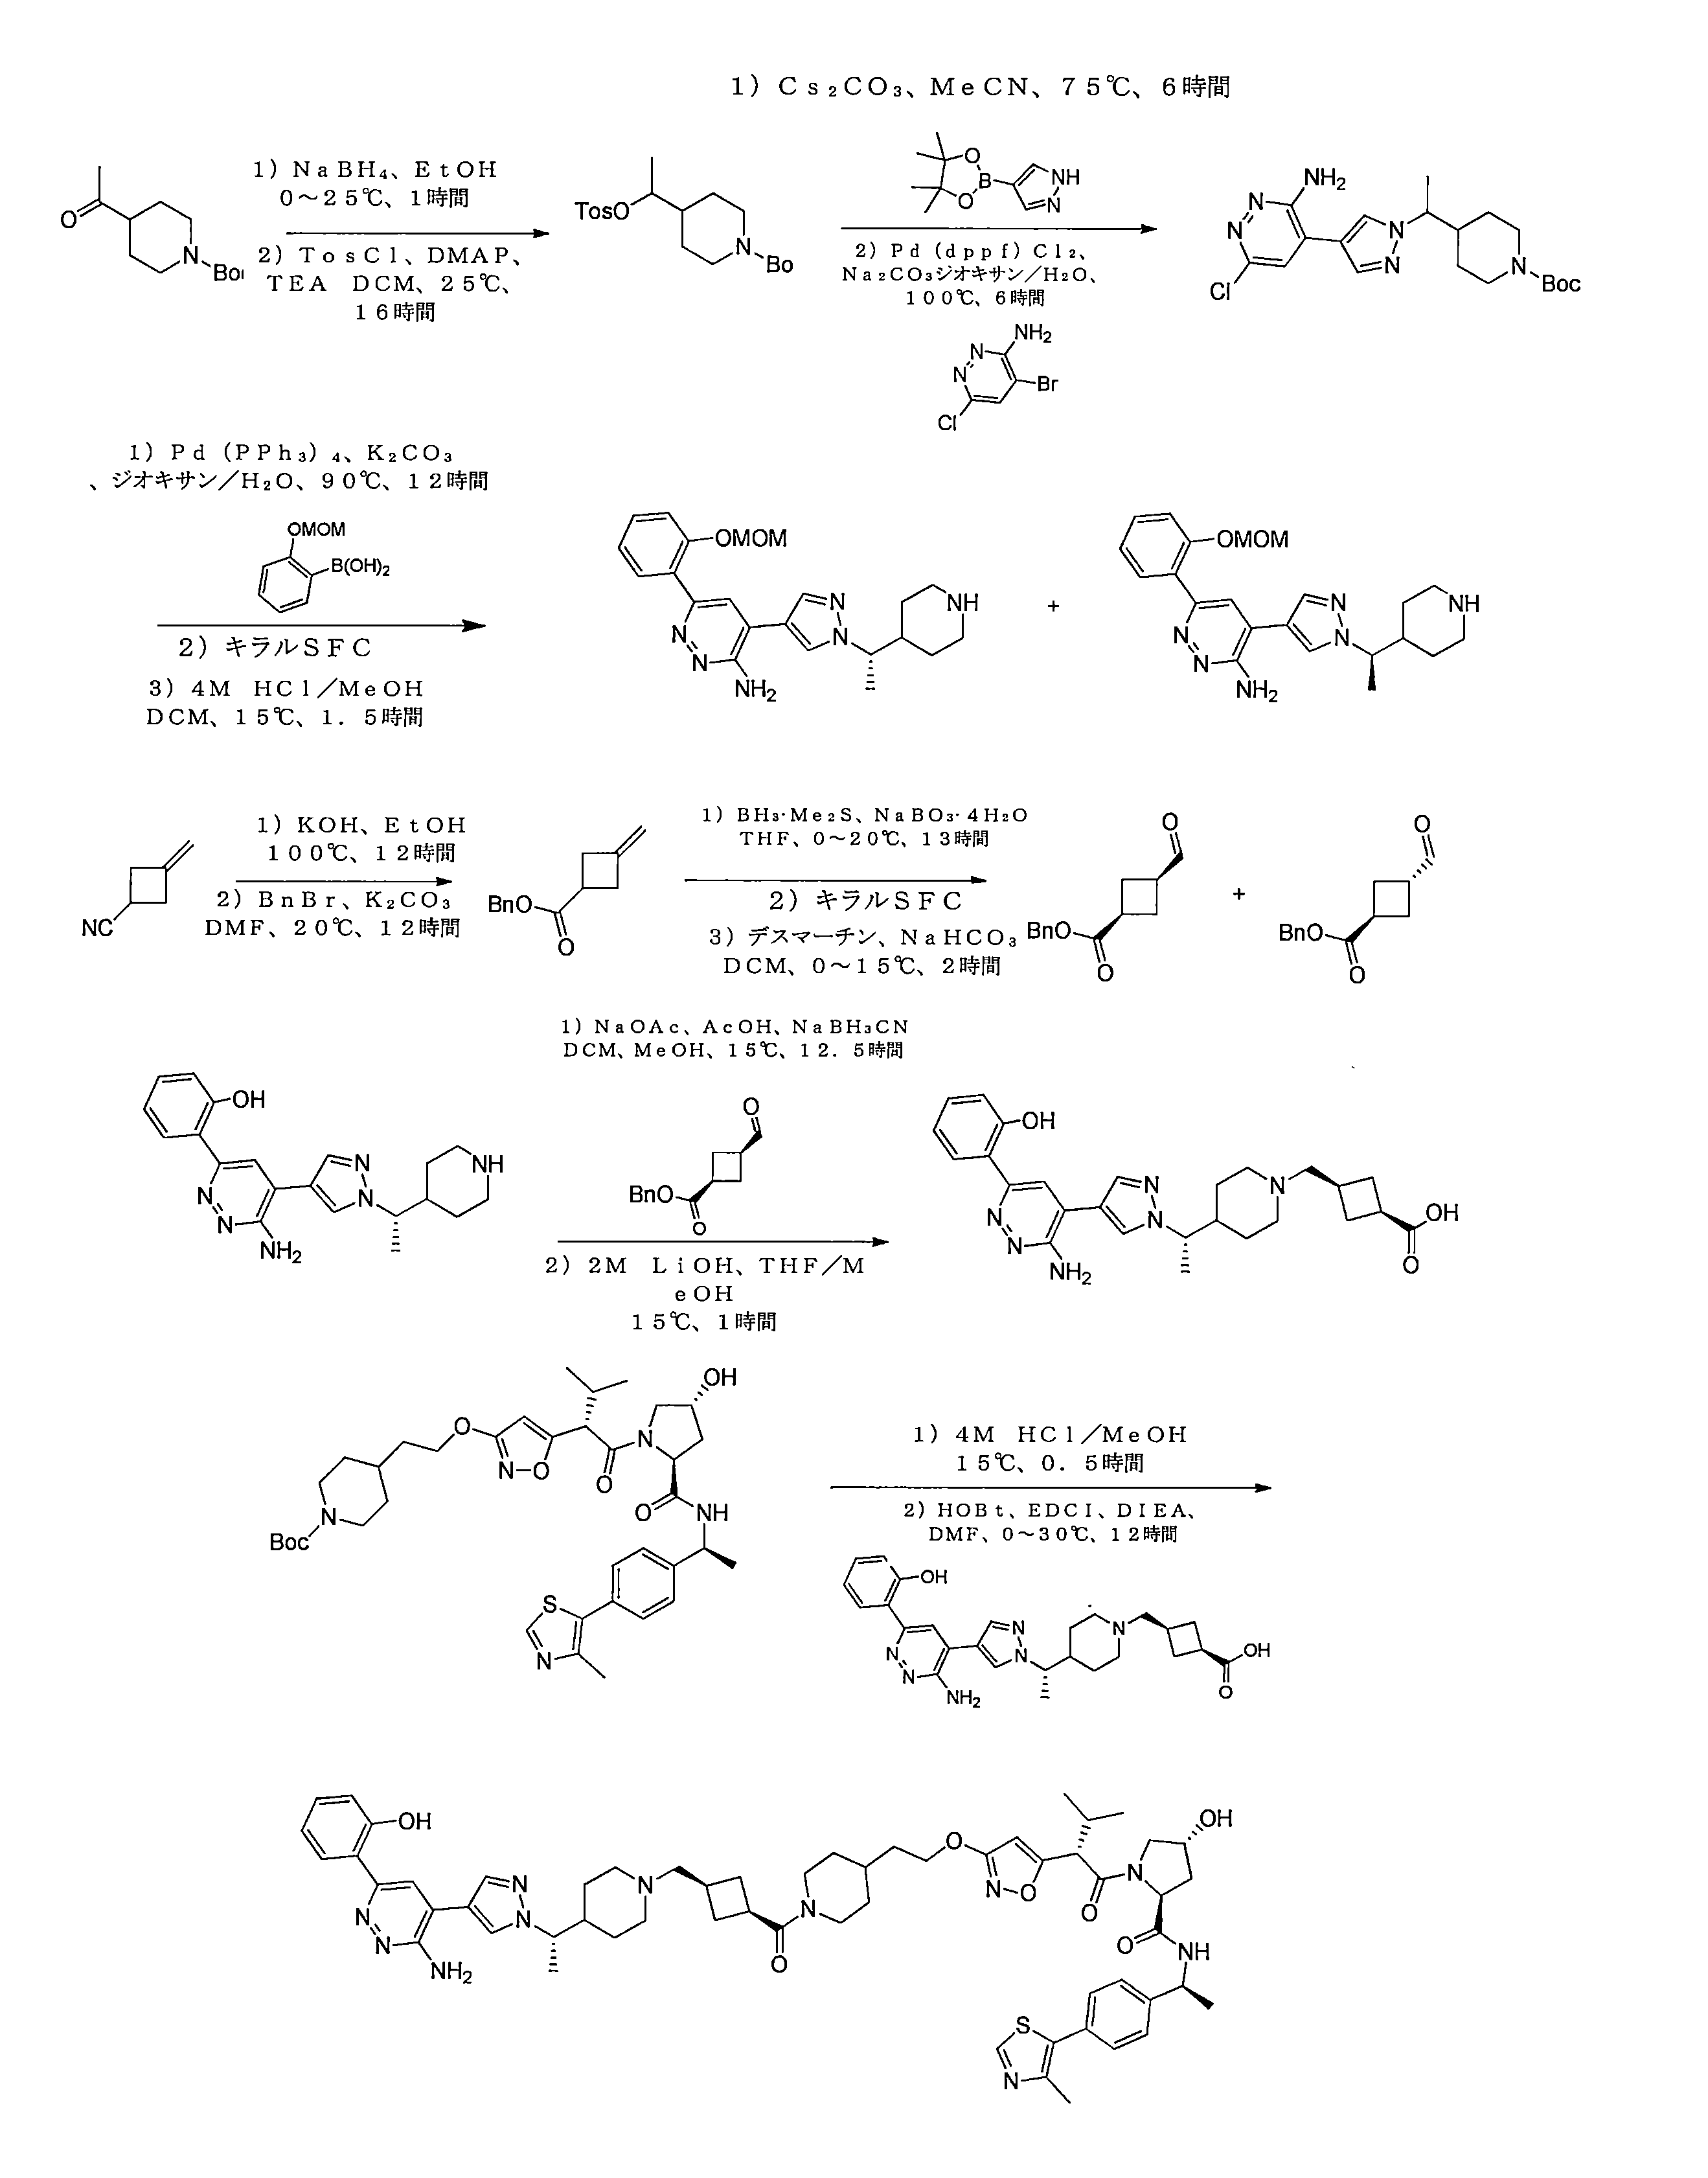
上記のスキームに示されるように、第1のカップリング断片のアセンブリは、tert-ブチル4-アセチルピペリジン-1-カルボキシレートで開始し、それをアルコールに還元し、トシレートに変換した。トシル基を、4-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)-1H-ピラゾールで置換し、生成物を、スズキ条件下で4-ブロモ-6-クロロピリダジン-3-アミンとカップリングして、tert-ブチル4-(1-(4-(3-アミノ-6-(2-(メトキシメトキシ)フェニル)ピリダジン-4-イル)-1H-ピラゾール-1-イル)エチル)ピペリジン-1-カルボキシレートのラセミ混合物を得、それをキラル超臨界流体クロマトグラフィー(SFC)を使用することによって分離した。その後のBoc脱保護は、鏡像異性的に純粋な状態で所望の第1のカップリング断片を提供した。 As shown in the scheme above, the assembly of the first coupling fragment started with tert-butyl 4-acetylpiperidine-1-carboxylate, which was reduced to the alcohol and converted to the tosylate. The tosyl group was replaced with 4-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)-1H-pyrazole, and the product was coupled with 4-bromo-6-chloropyridazin-3-amine under Suzuki conditions to give a racemic mixture of tert-butyl 4-(1-(4-(3-amino-6-(2-(methoxymethoxy)phenyl)pyridazin-4-yl)-1H-pyrazol-1-yl)ethyl)piperidine-1-carboxylate, which was separated by using chiral supercritical fluid chromatography (SFC). Subsequent Boc deprotection provided the desired first coupling fragment in enantiomerically pure form.
第2のカップリング断片の形成は、3-メチレンシクロブタンカルボニトリルで開始し、それをカルボン酸に加水分解し、ベンジルエステルとして保護した。エンド二重結合の加水分離、続いて酸化は、アルコールのジアステレオマー混合物を提供し、それを超臨界流体クロマトグラフィー(SFC)を使用することによって分離した。個々のアルコールを、デスマーチンペリオジナンで酸化して、所望の異性体アルデヒドを得た。 Formation of the second coupling fragment began with 3-methylenecyclobutanecarbonitrile, which was hydrolyzed to the carboxylic acid and protected as the benzyl ester. Hydrolytic cleavage of the endo double bond followed by oxidation provided a diastereomeric mixture of alcohols, which were separated by using supercritical fluid chromatography (SFC). The individual alcohols were oxidized with Dess-Martin periodinane to give the desired isomeric aldehydes.
水素化ホウ素ナトリウムによる、第1および第2のカップリング断片の還元的アミノ化、続いて塩基加水分解により、(1R,3S)-3-((4-((S)-1-(4-(3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル)-1H-ピラゾール-1-イル)エチル)ピペリジン-1-イル)メチル)シクロブタンカルボン酸を得、それを(2S,4R)-4-ヒドロキシ-1-((S)-3-メチル-2-(3-(2-(ピペリジン-4-イル)エトキシ)イソキサゾール-5-イル)ブタノイル)-N-((S)-1-(4-(4-メチルチアゾール-5-イル)フェニル)エチル)ピロリジン-2-カルボキサミドと縮合して、所望の生成物を得た。この方法は、適切な立体化学を有する試薬および中間体を導入することによって、所望の生成物のすべての鏡像異性的に純粋な異性体を合成するのに好都合に使用され得る。 Reductive amination of the first and second coupling fragments with sodium borohydride followed by base hydrolysis gave (1R,3S)-3-((4-((S)-1-(4-(3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl)-1H-pyrazol-1-yl)ethyl)piperidin-1-yl)methyl)cyclobutanecarboxylic acid, which was condensed with (2S,4R)-4-hydroxy-1-((S)-3-methyl-2-(3-(2-(piperidin-4-yl)ethoxy)isoxazol-5-yl)butanoyl)-N-((S)-1-(4-(4-methylthiazol-5-yl)phenyl)ethyl)pyrrolidine-2-carboxamide to give the desired product. This method can be conveniently used to synthesize all enantiomerically pure isomers of the desired product by introducing reagents and intermediates with the appropriate stereochemistry.
例示的な化合物458の例示的な合成

上記のスキームに示されるように、tert-ブチル4-ホルミルピペリジン-1-カルボキシレートは、(2S,4R)-4-ヒドロキシ-1-((S)-3-メチル-2-(3-(ピペラジン-1-イル)イソキサゾール-5-イル)ブタノイル)-N-((S)-1-(4-(4-メチルチアゾール-5-イル)フェニル)エチル)ピロリジン-2-カルボキサミドおよびシアノホウ素化ナトリウムによる還元的アミノ化を受け、Boc脱保護後に(2S,4R)-4-ヒドロキシ-1-((S)-3-メチル-2-(3-(4-(ピペリジン-4-イルメチル)ピペラジン-1-イル)イソキサゾール-5-イル)ブタノイル)-N-((S)-1-(4-(4-メチルチアゾール-5-イル)フェニル)エチル)ピロリジン-2-カルボキサミドを生成した。同様の条件下での(1R,3R)-3-((4-(3-(3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル)オキシ)シクロブタンカルバルデヒドによる、得られた化合物のその後の還元的アミノ化は、所望の最終生成物を提供した。 As shown in the scheme above, tert-butyl 4-formylpiperidine-1-carboxylate underwent reductive amination with (2S,4R)-4-hydroxy-1-((S)-3-methyl-2-(3-(piperazin-1-yl)isoxazol-5-yl)butanoyl)-N-((S)-1-(4-(4-methylthiazol-5-yl)phenyl)ethyl)pyrrolidine-2-carboxamide and sodium cyanoborate to generate (2S,4R)-4-hydroxy-1-((S)-3-methyl-2-(3-(4-(piperidin-4-ylmethyl)piperazin-1-yl)isoxazol-5-yl)butanoyl)-N-((S)-1-(4-(4-methylthiazol-5-yl)phenyl)ethyl)pyrrolidine-2-carboxamide after Boc deprotection. Subsequent reductive amination of the resulting compound with (1R,3R)-3-((4-(3-(3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl)-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl)oxy)cyclobutanecarbaldehyde under similar conditions provided the desired final product.
例示的な化合物474の例示的な合成

上記のスキームに示されるように、4-ヒドロキシピリジンおよびtert-ブチル4-ヒドロキシピペリジン-1-カルボキシレートの間のミツノブ反応は、tert-ブチル4-(ピリジン-4-イルオキシ)ピペリジン-1-カルボキシレートを提供し、それを、ピリジン窒素を臭化ベンジルでアルキル化することによって塩に変換した。その後、ピリジン環を2段階手順によって還元して、二重ピペリジン誘導体を形成し、それを、ヘテロ芳香族求核性置換条件下でメチル3-メチル-2-(3-(((ペルフルオロブチル)スルホニル)オキシ)イソキサゾール-5-イル)ブタノエートと反応させて、カップリング生成物を形成した。エステル加水分解、続いて(2S,4R)-4-ヒドロキシ-N-((S)-1-(4-(4-メチルチアゾール-5-イル)フェニル)エチル)ピロリジン-2-カルボキサミドとの得られた酸の縮合は、ジアステレオマー最終生成物を提供し、それを超臨界流体クロマトグラフィー(SFC)によって分離した。 As shown in the scheme above, Mitsunobu reaction between 4-hydroxypyridine and tert-butyl 4-hydroxypiperidine-1-carboxylate provided tert-butyl 4-(pyridin-4-yloxy)piperidine-1-carboxylate, which was converted to a salt by alkylating the pyridine nitrogen with benzyl bromide. The pyridine ring was then reduced by a two-step procedure to form a double piperidine derivative, which was reacted with methyl 3-methyl-2-(3-(((perfluorobutyl)sulfonyl)oxy)isoxazol-5-yl)butanoate under heteroaromatic nucleophilic substitution conditions to form the coupling product. Ester hydrolysis followed by condensation of the resulting acid with (2S,4R)-4-hydroxy-N-((S)-1-(4-(4-methylthiazol-5-yl)phenyl)ethyl)pyrrolidine-2-carboxamide provided the diastereomeric final products, which were separated by supercritical fluid chromatography (SFC).
例示的な化合物494、495の例示的な合成

上記のスキームに示されるように、PTM結合部分の合成は、3-((4-メトキシベンジル)オキシ)プロパン-1-オールで開始し、それを水素化ナトリウムで処理して、(S)-2-メチルオキシランを開いた。得られたアルコールの臭素化、続いて2-(3-メチルイソキサゾール-5-イル)酢酸メチルに由来するエノレートによる臭化物置換は、(3R)-メチル4-(3-ヒドロキシプロポキシ)-3-メチル-2-(3-メチルイソキサゾール-5-イル)ブタノエートを提供した。4-ブロモピリジン-2-オールとの生成物のミツノブ反応、続いてモノ-Boc保護された3,8-ジアザビシクロ[3.2.1]オクタンとの臭化ピリジル部分のパラジウム触媒カップリングにより、tert-ブチル8-(2-(3-((2R)-4-メトキシ-2-メチル-3-(3-メチルイソキサゾール-5-イル)-4-オキソブトキシ)プロポキシ)ピリジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-3-カルボキシレートを得た。その後のBoc基切断は、遊離アミンを提供し、それはアミン(DIEA)の存在下で、非プロトン性極性溶媒(DMSO)中で加熱することによって、4-ブロモ-6-クロロピリダジン-3-アミンとの求核カップリング反応を受けた。ピリダジン臭化物と(2-ヒドロキシフェニル)ボロン酸とのスズキカップリング反応は、PTM結合部分のアセンブリを完了した。エステル加水分解、続いて(2S,4R)-4-ヒドロキシ-N-((S)-1-(4-(4-メチルチアゾール-5-イル)フェニル)エチル)ピロリジン-2-カルボキサミドとの縮合により、ジアステレオマーの混合物として所望の最終生成物を得、それを超臨界流体クロマトグラフィー(SFC)によって分離した。 As shown in the scheme above, the synthesis of the PTM binding moiety began with 3-((4-methoxybenzyl)oxy)propan-1-ol, which was treated with sodium hydride to open the (S)-2-methyloxirane. Bromination of the resulting alcohol followed by bromide displacement with the enolate derived from methyl 2-(3-methylisoxazol-5-yl)acetate provided (3R)-methyl 4-(3-hydroxypropoxy)-3-methyl-2-(3-methylisoxazol-5-yl)butanoate. Mitsunobu reaction of the product with 4-bromopyridin-2-ol followed by palladium catalyzed coupling of the pyridyl bromide moiety with mono-Boc protected 3,8-diazabicyclo[3.2.1]octane gave tert-butyl 8-(2-(3-((2R)-4-methoxy-2-methyl-3-(3-methylisoxazol-5-yl)-4-oxobutoxy)propoxy)pyridin-4-yl)-3,8-diazabicyclo[3.2.1]octane-3-carboxylate. Subsequent Boc group cleavage provided the free amine, which underwent nucleophilic coupling reaction with 4-bromo-6-chloropyridazin-3-amine by heating in aprotic polar solvent (DMSO) in the presence of amine (DIEA). Suzuki coupling reaction of the pyridazine bromide with (2-hydroxyphenyl)boronic acid completed the assembly of the PTM binding moiety. Ester hydrolysis followed by condensation with (2S,4R)-4-hydroxy-N-((S)-1-(4-(4-methylthiazol-5-yl)phenyl)ethyl)pyrrolidine-2-carboxamide gave the desired final products as a mixture of diastereomers, which were separated by supercritical fluid chromatography (SFC).
例示的な化合物496、497の例示的な合成

上記のスキームに示されるように、所望の生成物を、シアノ水素化ホウ素ナトリウムの存在下で、2-(6-アミノ-5-(1-((R)-4-((1r,3R)-3-(ピペリジン-4-イルオキシ)シクロブチル)ブタ-3-イン-2-イル)-1H-ピラゾール-4-イル)ピリダジン-3-イル)フェノールによる、(2S,4R)-1-((R)-2-(3-(4-ホルミルピペリジン-1-イル)イソキサゾール-5-イル)-3-メチルブタノイル)-4-ヒドロキシ-N-((S)-1-(4-(4-メチルチアゾール-5-イル)フェニル)エチル)ピロリジン-2-カルボキサミドの還元的アミノ化によって得た。この方法は、適切な立体化学を有するカップリング対応物を導入することによって、所望の生成物のすべての鏡像異性的に純粋な異性体を合成するのに好都合に使用され得る。 As shown in the above scheme, the desired product was obtained by reductive amination of (2S,4R)-1-((R)-2-(3-(4-formylpiperidin-1-yl)isoxazol-5-yl)-3-methylbutanoyl)-4-hydroxy-N-((S)-1-(4-(4-methylthiazol-5-yl)phenyl)ethyl)pyrrolidine-2-carboxamide with 2-(6-amino-5-(1-((R)-4-((1r,3R)-3-(piperidin-4-yloxy)cyclobutyl)but-3-yn-2-yl)-1H-pyrazol-4-yl)pyridazin-3-yl)phenol in the presence of sodium cyanoborohydride. This method can be conveniently used to synthesize all enantiomerically pure isomers of the desired product by introducing the coupling counterpart with the appropriate stereochemistry.
例示的な化合物499の例示的な合成
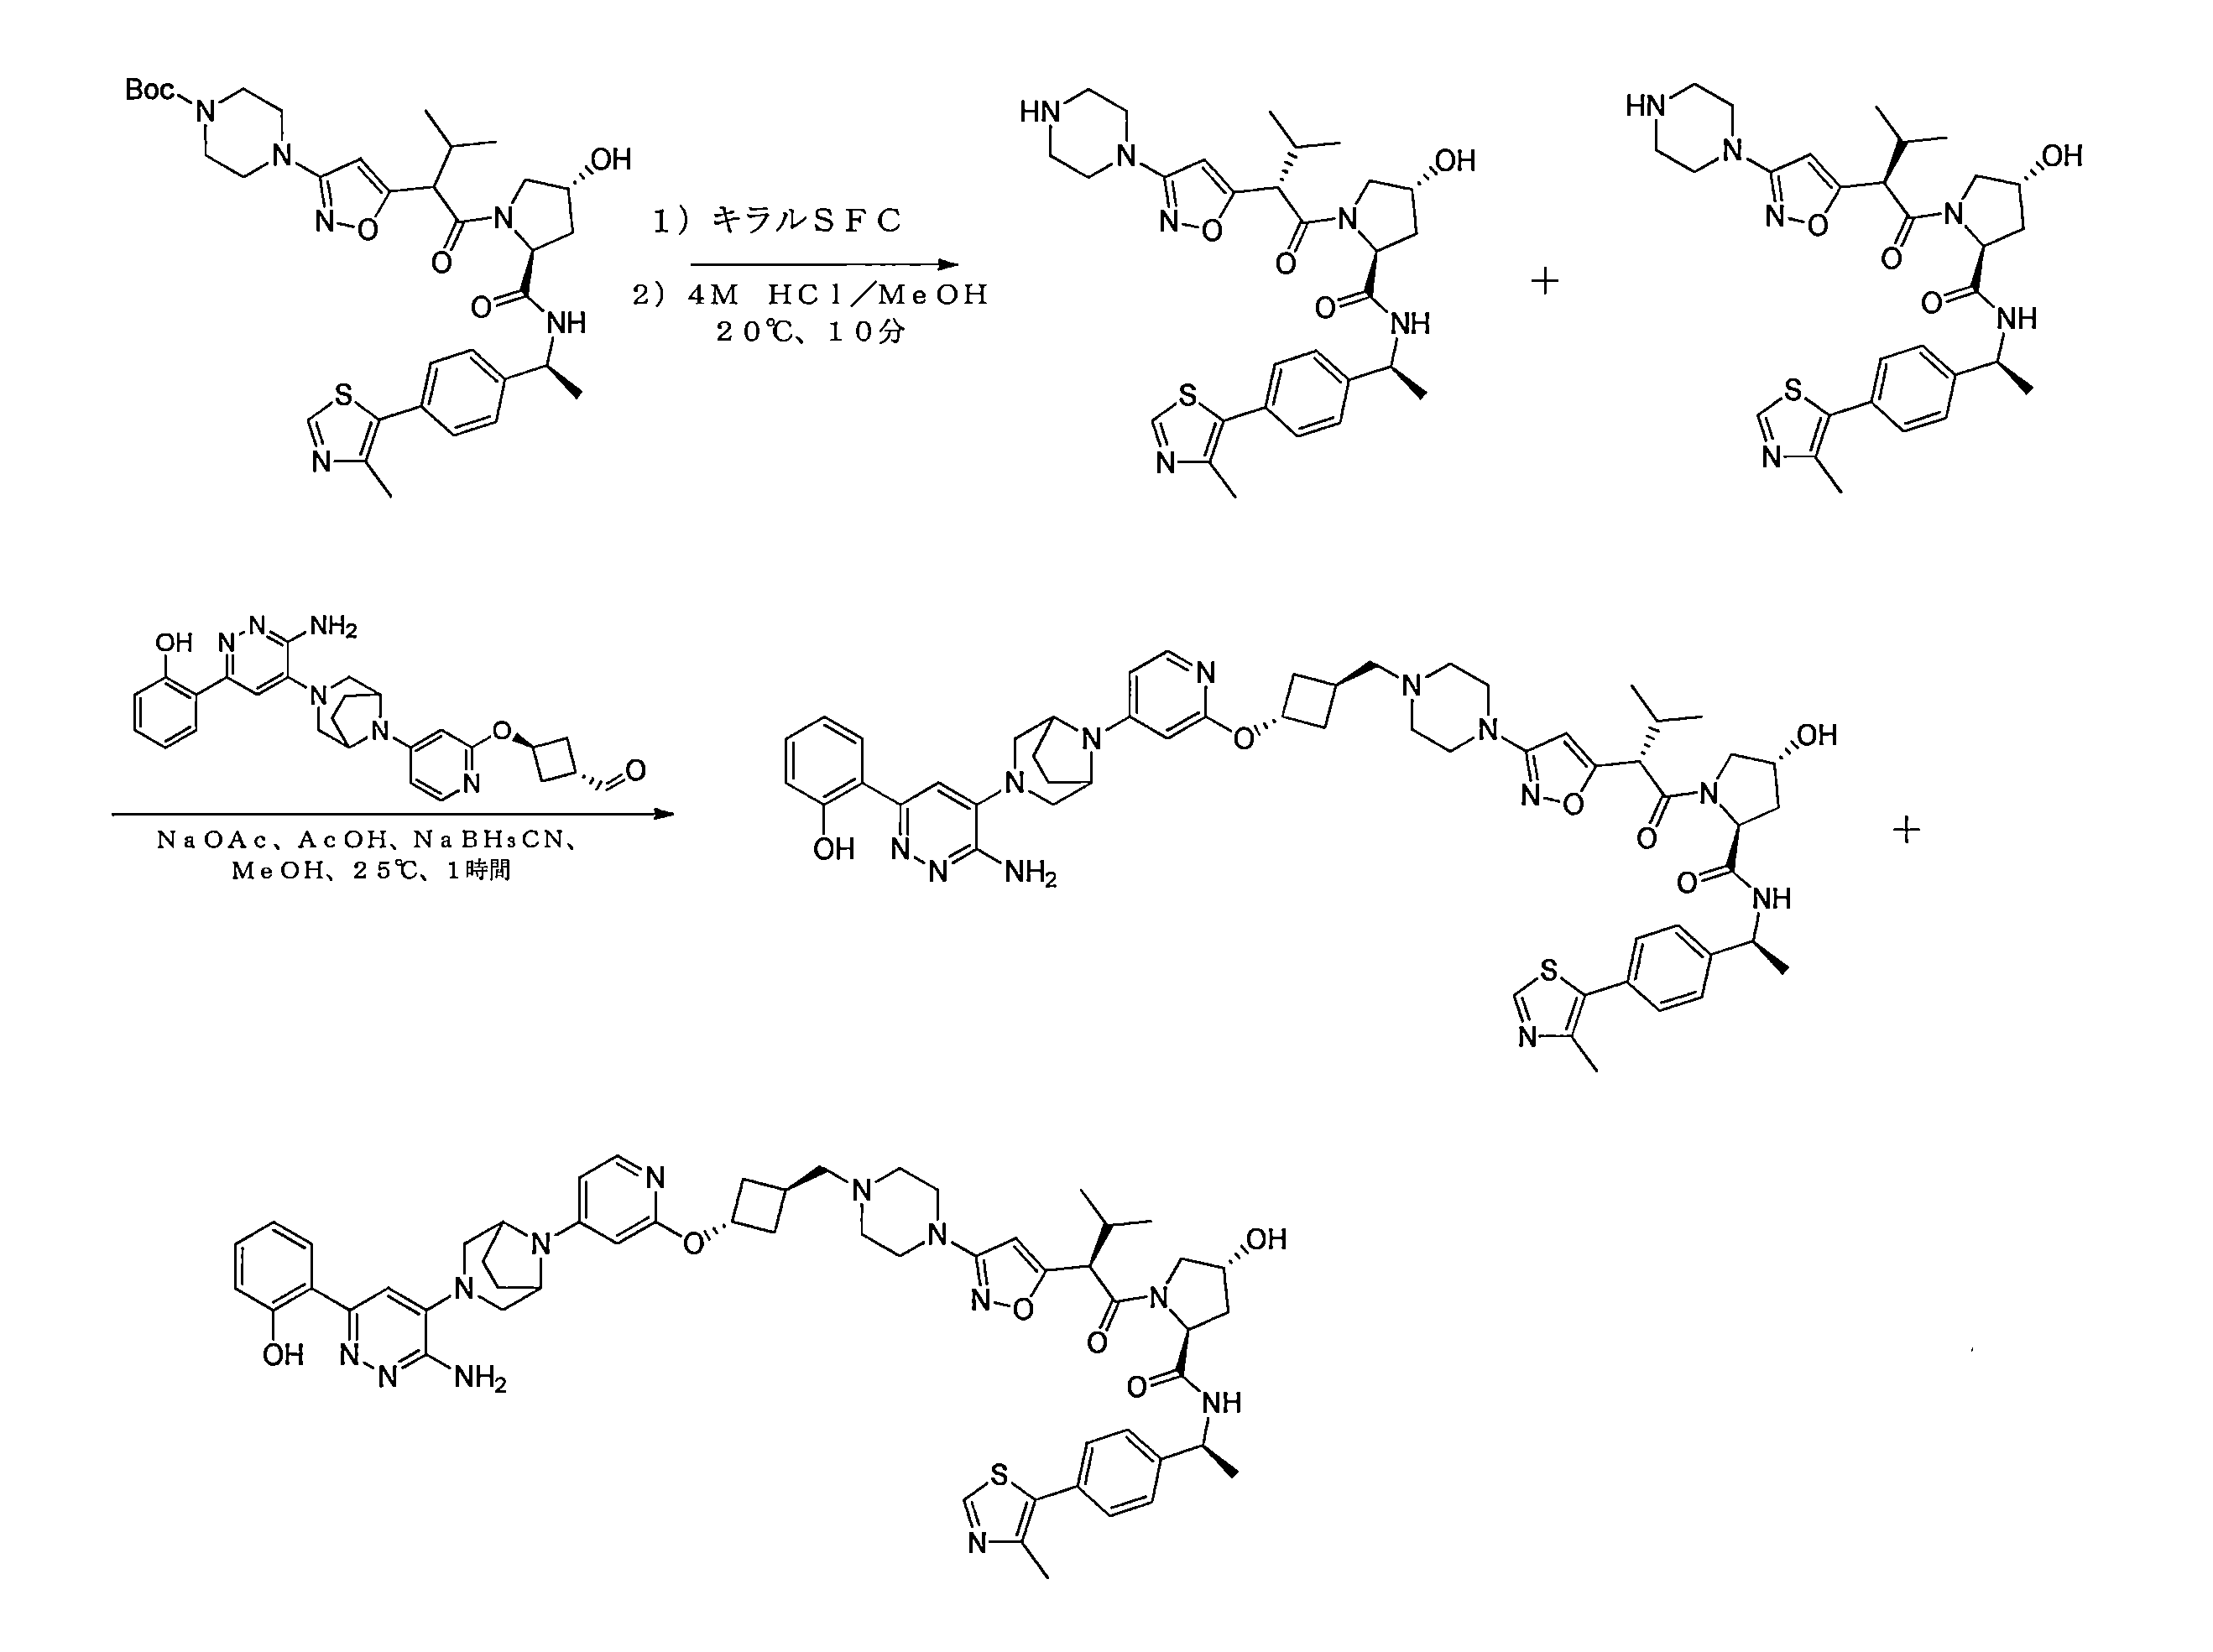
上記のスキームに示されるように、ジアステレオマー混合物としてのtert-ブチル4-(5-(1-((2S,4R)-4-ヒドロキシ-2-(((S)-1-(4-(4-メチルチアゾール-5-イル)フェニル)エチル)カルバモイル)ピロリジン-1-イル)-3-メチル-1-オキソブタン-2-イル)イソキサゾール-3-イル)ピペラジン-1-カルボキシレートを、個々の異性体に分離した。次いで、これらの異性体の各々を、(1R,3R)-3-((4-(3-(3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル)-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル)ピリジン-2-イル)オキシ)シクロブタンカルバルデヒドと反応させて、鏡像異性的に純粋な形態で所望の最終生成物を提供した。この方法は、適切な立体化学を有するカップリング対応物を導入することによって、所望の生成物のすべての鏡像異性的に純粋な異性体を合成するのに好都合に使用され得る。 As shown in the scheme above, tert-butyl 4-(5-(1-((2S,4R)-4-hydroxy-2-(((S)-1-(4-(4-methylthiazol-5-yl)phenyl)ethyl)carbamoyl)pyrrolidin-1-yl)-3-methyl-1-oxobutan-2-yl)isoxazol-3-yl)piperazine-1-carboxylate as a diastereomeric mixture was separated into individual isomers. Each of these isomers was then reacted with (1R,3R)-3-((4-(3-(3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl)-3,8-diazabicyclo[3.2.1]octan-8-yl)pyridin-2-yl)oxy)cyclobutanecarbaldehyde to provide the desired final product in enantiomerically pure form. This method can be conveniently used to synthesize all enantiomerically pure isomers of a desired product by introducing coupling counterparts with the appropriate stereochemistry.
例示的な化合物143の例示的な合成
工程1
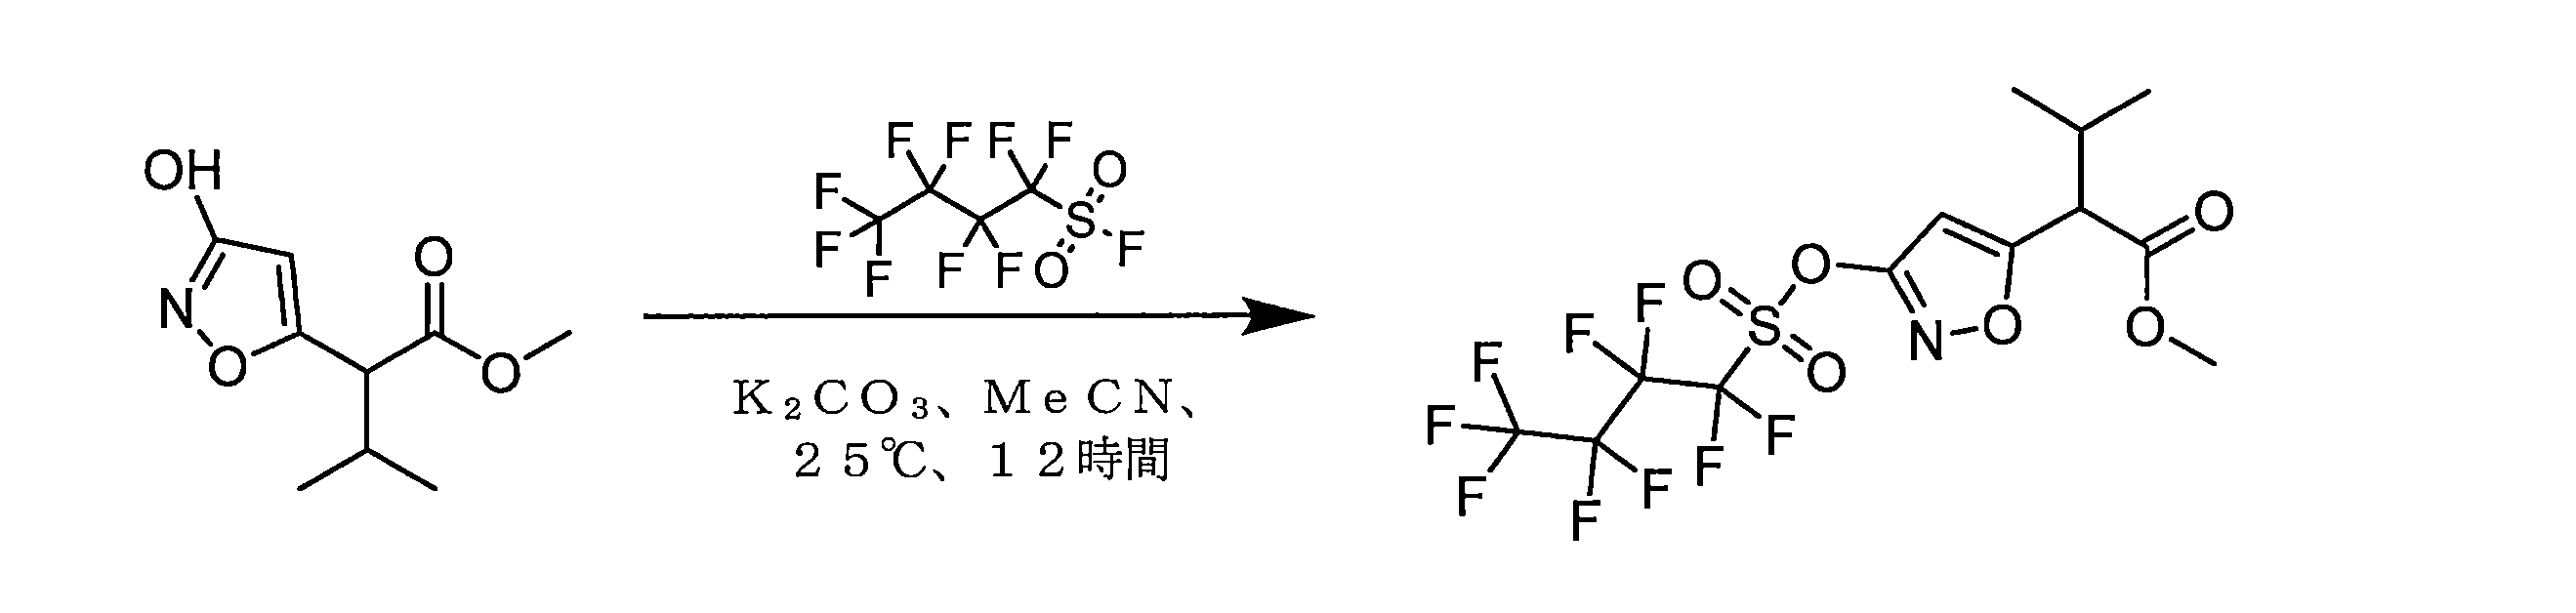
MeCN(1.6L)中の2-(3-メチルイソキサゾール-5-イル)-3-メチルブタン酸メチル(265g、1.33mol)[例示的な化合物144について記載されるように調製した〕の溶液に、K2CO3(367.71g、2.66mol)および1,1,2,2,3,3,4,4,4-ノナフルオロブタン-1-スルホニルフルオリド(482.25g、1.60mol、280.38mL)を添加した。反応混合物を、25℃で12時間撹拌した。反応混合物を減圧下で濃縮し、MeCNを除去した。残渣を水(1.5L)で希釈し、酢酸エチル(500mL×3)で抽出した。合わせた有機層を、ブライン(200mL)で洗浄し、Na2SO4上で乾燥させ、濾過し、減圧下で濃縮して、3-メチル-2-(3-(((ペルフルオロブチル)スルホニル)オキシ)イソキサゾール-5-イル)ブタン酸メチル(458g、886.42mmol)を茶色の油として得た。 To a solution of methyl 2-(3-methylisoxazol-5-yl)-3-methylbutanoate (265 g, 1.33 mol) [prepared as described for exemplary compound 144] in MeCN (1.6 L), K2CO3 (367.71 g, 2.66 mol) and 1,1,2,2,3,3,4,4,4-nonafluorobutane-1-sulfonyl fluoride (482.25 g, 1.60 mol, 280.38 mL) were added. The reaction mixture was stirred at 25 °C for 12 h. The reaction mixture was concentrated under reduced pressure to remove MeCN. The residue was diluted with water (1.5 L) and extracted with ethyl acetate (500 mL x 3). The combined organic layers were washed with brine (200 mL), dried over Na2SO4, filtered, and concentrated under reduced pressure to give methyl 3-methyl-2-(3-(((perfluorobutyl)sulfonyl)oxy)isoxazol-5-yl)butanoate (458 g, 886.42 mmol) as a brown oil.
工程2

DMF(1.6L)中の3-メチル-2-(3-(((ペルフルオロブチル)スルホニル)オキシ)イソキサゾール-5-イル)ブタン酸メチル(160g、332.44mmol)および4-(ジメトキシメチル)ピペリジン(79.40g、498.66mmol)の混合物に、DIEA(128.90g、997.32mmol、173.72mL)を、窒素下で25℃において一度に添加した。混合物を80℃で12時間撹拌した。混合物を25℃に冷却し、氷水(w:w=1:1)(3L)に注ぎ入れ、15分間撹拌した。水相を酢酸エチル(1L×5)で抽出した。合わせた有機相をブライン(300mL×3)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、真空中で濃縮した。混合物をEtOAc(70mL)中に溶解し、濾過して、不溶性材料を除去した。残渣を、カラムクロマトグラフィー(石油エーテル/酢酸エチル=80:1~0:1)によって精製して、(34g、100mmol)の2-(3-(4-(ジメトキシメチル)ピペリジン-1-イル)イソキサゾール-5-イル)-3-メチルブタン酸メチルをオレンジ色のシロップとして得た。 To a mixture of methyl 3-methyl-2-(3-(((perfluorobutyl)sulfonyl)oxy)isoxazol-5-yl)butanoate (160 g, 332.44 mmol) and 4-(dimethoxymethyl)piperidine (79.40 g, 498.66 mmol) in DMF (1.6 L) was added DIEA (128.90 g, 997.32 mmol, 173.72 mL) in one portion at 25° C. under nitrogen. The mixture was stirred at 80° C. for 12 h. The mixture was cooled to 25° C. and poured into ice water (w:w=1:1) (3 L) and stirred for 15 min. The aqueous phase was extracted with ethyl acetate (1 L×5). The combined organic phase was washed with brine (300 mL×3), dried over anhydrous sodium sulfate, filtered and concentrated in vacuo. The mixture was dissolved in EtOAc (70 mL) and filtered to remove insoluble material. The residue was purified by column chromatography (petroleum ether/ethyl acetate = 80:1 to 0:1) to obtain methyl 2-(3-(4-(dimethoxymethyl)piperidin-1-yl)isoxazol-5-yl)-3-methylbutanoate (34 g, 100 mmol) as an orange syrup.
工程3

テトラヒドロフラン(1mL)およびメタノール(1mL)中の2-[3-[4-(ジメトキシメチル)-1-ピペリジル]イソキサゾール-5-イル]-3-メチル-ブタン酸メチル(350mg、1.03mmol、1当量)の混合物に、水酸化リチウム(2M、1mL、1.95当量)を窒素下で一度に添加した。混合物を25℃で1時間撹拌した。トリフルオロ酢酸を用いてpHを5~6に調整し、45℃において減圧下で濃縮した。混合物を、分取HPLC(カラム:Phenomenex Synergi C18 150*25*10um;移動相:[水(0.225%FA)-ACN];B%:40%~70%、9分)によって精製して、2-[3-[4-(ジメトキシメチル)-1-ピペリジル]イソキサゾール-5-イル]-3-メチル-ブタン酸(190mg)を黄色の油として得た。 To a mixture of methyl 2-[3-[4-(dimethoxymethyl)-1-piperidyl]isoxazol-5-yl]-3-methyl-butanoate (350 mg, 1.03 mmol, 1 equiv.) in tetrahydrofuran (1 mL) and methanol (1 mL), lithium hydroxide (2 M, 1 mL, 1.95 equiv.) was added in one portion under nitrogen. The mixture was stirred at 25°C for 1 h. The pH was adjusted to 5-6 with trifluoroacetic acid and concentrated under reduced pressure at 45°C. The mixture was purified by preparative HPLC (column: Phenomenex Synergi C18 150*25*10um; mobile phase: [water (0.225% FA)-ACN]; B%: 40%-70%, 9 min) to give 2-[3-[4-(dimethoxymethyl)-1-piperidyl]isoxazol-5-yl]-3-methyl-butanoic acid (190 mg) as a yellow oil.
工程4

N,N-ジメチルホルムアミド(10mL)中の2-[3-[4-(ジメトキシメチル)-1-ピペリジル]イソキサゾール-5-イル]-3-メチル-ブタン酸(190mg、0.58mmol、1当量)、N,N-ジイソプロピルエチルアミン(225mg、1.75mmol、0.3mL、3当量)、および(2S,4R)-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(257mg、0.70mmol、1.2当量、塩酸塩)の混合物に、O-(7-アザベンゾトリアゾール-1-イル)-N,N,N,N-テトラメチルウロニウムヘキサフルオロホスフェート(288mg、0.76mmol、1.3当量)を、窒素下で0℃において一度に添加した。混合物を25℃で1時間撹拌した。混合物を氷水(w/w=1/1)(30mL)に注ぎ入れ、15分間撹拌した。水相を酢酸エチル(30mL×3)で抽出した。合わせた有機層をブライン(30mL×3)で洗浄し、硫酸ナトリウムで乾燥させ、濾過し、真空中で濃縮した。残渣を、シリカゲルクロマトグラフィー(ジクロロメタン:メタノール=10/1)によって精製して、(2S,4R)-1-[2-[3-[4-(ジメトキシメチル)-1-ピペリジル]イソキサゾール-5-イル]-3-メチル-ブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(208mg)を黄色の油として得た。 To a mixture of 2-[3-[4-(dimethoxymethyl)-1-piperidyl]isoxazol-5-yl]-3-methyl-butanoic acid (190 mg, 0.58 mmol, 1 eq.), N,N-diisopropylethylamine (225 mg, 1.75 mmol, 0.3 mL, 3 eq.), and (2S,4R)-4-hydroxy-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (257 mg, 0.70 mmol, 1.2 eq., hydrochloride salt) in N,N-dimethylformamide (10 mL) was added O-(7-azabenzotriazol-1-yl)-N,N,N,N-tetramethyluronium hexafluorophosphate (288 mg, 0.76 mmol, 1.3 eq.) in one portion at 0°C under nitrogen. The mixture was stirred at 25°C for 1 hour. The mixture was poured into ice water (w/w=1/1) (30 mL) and stirred for 15 minutes. The aqueous phase was extracted with ethyl acetate (30 mL x 3). The combined organic layers were washed with brine (30 mL x 3), dried over sodium sulfate, filtered, and concentrated in vacuo. The residue was purified by silica gel chromatography (dichloromethane:methanol=10/1) to give (2S,4R)-1-[2-[3-[4-(dimethoxymethyl)-1-piperidyl]isoxazol-5-yl]-3-methyl-butanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (208 mg) as a yellow oil.
工程5

(2S,4R)-1-[2-[3-[4-(ジメトキシメチル)-1-ピペリジル]イソキサゾール-5-イル]-3-チル-ブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(200mg、0.31mmol、1当量)を、SFC(カラム:DAICEL CHIRALPAK AD(250mm*30mm、10um);移動相:[0.1%NH3H2O IPA];B%:45%~45%、3.0分;65分)によって精製した。混合物を、減圧下で、45℃で濃縮した。 (2S,4R)-1-[2-[3-[4-(dimethoxymethyl)-1-piperidyl]isoxazol-5-yl]-3-thyl-butanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (200 mg, 0.31 mmol, 1 eq.) was purified by SFC (column: DAICEL CHIRALPAK AD (250 mm*30 mm, 10 um); mobile phase: [0.1% NH3H2O IPA]; B%: 45%-45%, 3.0 min; 65 min). The mixture was concentrated under reduced pressure at 45°C.
(2S,4R)-1-[(2S)-2-[3-[4-(ジメトキシメチル)-1-ピペリジル]イソキサゾール-5-イル]-3-メチル-ブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(90mg)(保持時間:キラルSFCにおいて1.884分)を黄色の油として得た。 (2S,4R)-1-[(2S)-2-[3-[4-(dimethoxymethyl)-1-piperidyl]isoxazol-5-yl]-3-methyl-butanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (90 mg) (retention time: 1.884 min in chiral SFC) was obtained as a yellow oil.
(2S,4R)-1-[(2S)-2-[3-[4-(ジメトキシメチル)-1-ピペリジル]イソキサゾール-5-イル]-3-メチル-ブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(90mg、0.13mmol、収率42%、純度94%)(保持時間:キラルSFCにおいて2.230分)を黄色の油として得た。 (2S,4R)-1-[(2S)-2-[3-[4-(dimethoxymethyl)-1-piperidyl]isoxazol-5-yl]-3-methyl-butanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (90 mg, 0.13 mmol, 42% yield, 94% purity) (retention time: 2.230 min in chiral SFC) was obtained as a yellow oil.
工程6

テトラヒドロフラン(2mL)中の(2S,4R)-1-[(2R)-2-[3-[4-ジメトキシメチル)-1-ピペリジル]イソキサゾール-5-イル]-3-メチル-ブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(80mg、0.13mmol、1当量)の溶液に、硫酸溶液(1M、2mL、16.00当量)を添加した。反応混合物を50℃で5時間撹拌した。混合物を飽和重炭酸ナトリウム(10mL)に添加し、混合物を酢酸エチル(20mL×3)で抽出した。合わせた有機相をブライン(30mL)で洗浄し、硫酸ナトリウム上で乾燥させ、濾過し、真空中で濃縮し、次の工程で直接使用した。粗(2S,4R)-1-[(2R)-2-[3-(4-ホルミル-1-ピペリジル)イソキサゾール-5-イル]-3-メチル-ブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(68mg)を淡い黄色の固体として得、次の工程で直接使用した。 To a solution of (2S,4R)-1-[(2R)-2-[3-[4-dimethoxymethyl)-1-piperidyl]isoxazol-5-yl]-3-methyl-butanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (80 mg, 0.13 mmol, 1 equiv.) in tetrahydrofuran (2 mL), sulfuric acid solution (1 M, 2 mL, 16.00 equiv.) was added. The reaction mixture was stirred at 50 °C for 5 h. The mixture was added to saturated sodium bicarbonate (10 mL) and the mixture was extracted with ethyl acetate (20 mL x 3). The combined organic phase was washed with brine (30 mL), dried over sodium sulfate, filtered, concentrated in vacuo, and used directly in the next step. Crude (2S,4R)-1-[(2R)-2-[3-(4-formyl-1-piperidyl)isoxazol-5-yl]-3-methyl-butanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (68 mg) was obtained as a pale yellow solid and used directly in the next step.
工程7

メタノール(40mL)中の2-[6-アミノ-5-[8-[2-[3-(4-ピペリジルオキシ)シクロブトキシ]-4-ピリジル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-イル]ピリダジン-3-イル]フェノール二塩酸塩(2.08g、3.37mmol、1当量)の溶液に、酢酸ナトリウム(829mg、10.11mmol、3当量)を添加し、次いで、混合物に、(2S,4R)-1-[(2R)-2-[3-(4-ホルミル-1-ピペリジル)イソキサゾール-5-イル]-3-メチル-ブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(2g、3.37mmol、1当量)および酢酸(23mg、0.33mmol、0.1当量)を添加した。混合物を25℃で0.5時間撹拌し、次いで、混合物に、シアノホウ化水素ナトリウム(1.06g、16.84mmol、5当量)を添加した。混合物を25℃で2時間撹拌した。反応混合物を減圧下で濃縮した。残渣を、分取HPLC(カラム:Phenomenex luna C18 250*50mm*10um;移動相:[水(0.225%FA)-ACN];B%:10%~40%、22分)によって精製した。(2S,4R)-1-[(2R)-2-[3-[4-[[4-[3-[[4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]-2-ピリジル]オキシ]シクロブトキシ]-1-ピペリジル]メチル]-1-ピペリジル]イソキサゾール-5-イル]-3-メチル-ブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミドギ酸塩(1196mg、1.02mmol)を白色の固体として得た。 To a solution of 2-[6-amino-5-[8-[2-[3-(4-piperidyloxy)cyclobutoxy]-4-pyridyl]-3,8-diazabicyclo[3.2.1]octan-3-yl]pyridazin-3-yl]phenol dihydrochloride (2.08 g, 3.37 mmol, 1 eq.) in methanol (40 mL), sodium acetate (829 mg, 10.11 mmol, 3 eq.) was added, and the mixture was then cooled to 100° C. To the mixture was added (2S,4R)-1-[(2R)-2-[3-(4-formyl-1-piperidyl)isoxazol-5-yl]-3-methyl-butanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (2 g, 3.37 mmol, 1 eq.) and acetic acid (23 mg, 0.33 mmol, 0.1 eq.). The mixture was stirred at 25° C. for 0.5 h, and then to the mixture was added sodium cyanoborohydride (1.06 g, 16.84 mmol, 5 eq.). The mixture was stirred at 25° C. for 2 h. The reaction mixture was concentrated under reduced pressure. The residue was purified by preparative HPLC (column: Phenomenex luna C18 250*50mm*10um; mobile phase: [water (0.225% FA)-ACN]; B%: 10%-40%, 22 min). (2S,4R)-1-[(2R)-2-[3-[4-[[4-[3-[[4-[3-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl]-2-pyridyl]oxy]cyclobutoxy]-1-piperidyl]methyl]-1-piperidyl]isoxazol-5-yl]-3-methyl-butanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide formate (1196 mg, 1.02 mmol) was obtained as a white solid.
例示的な化合物143の例示的な合成
工程1
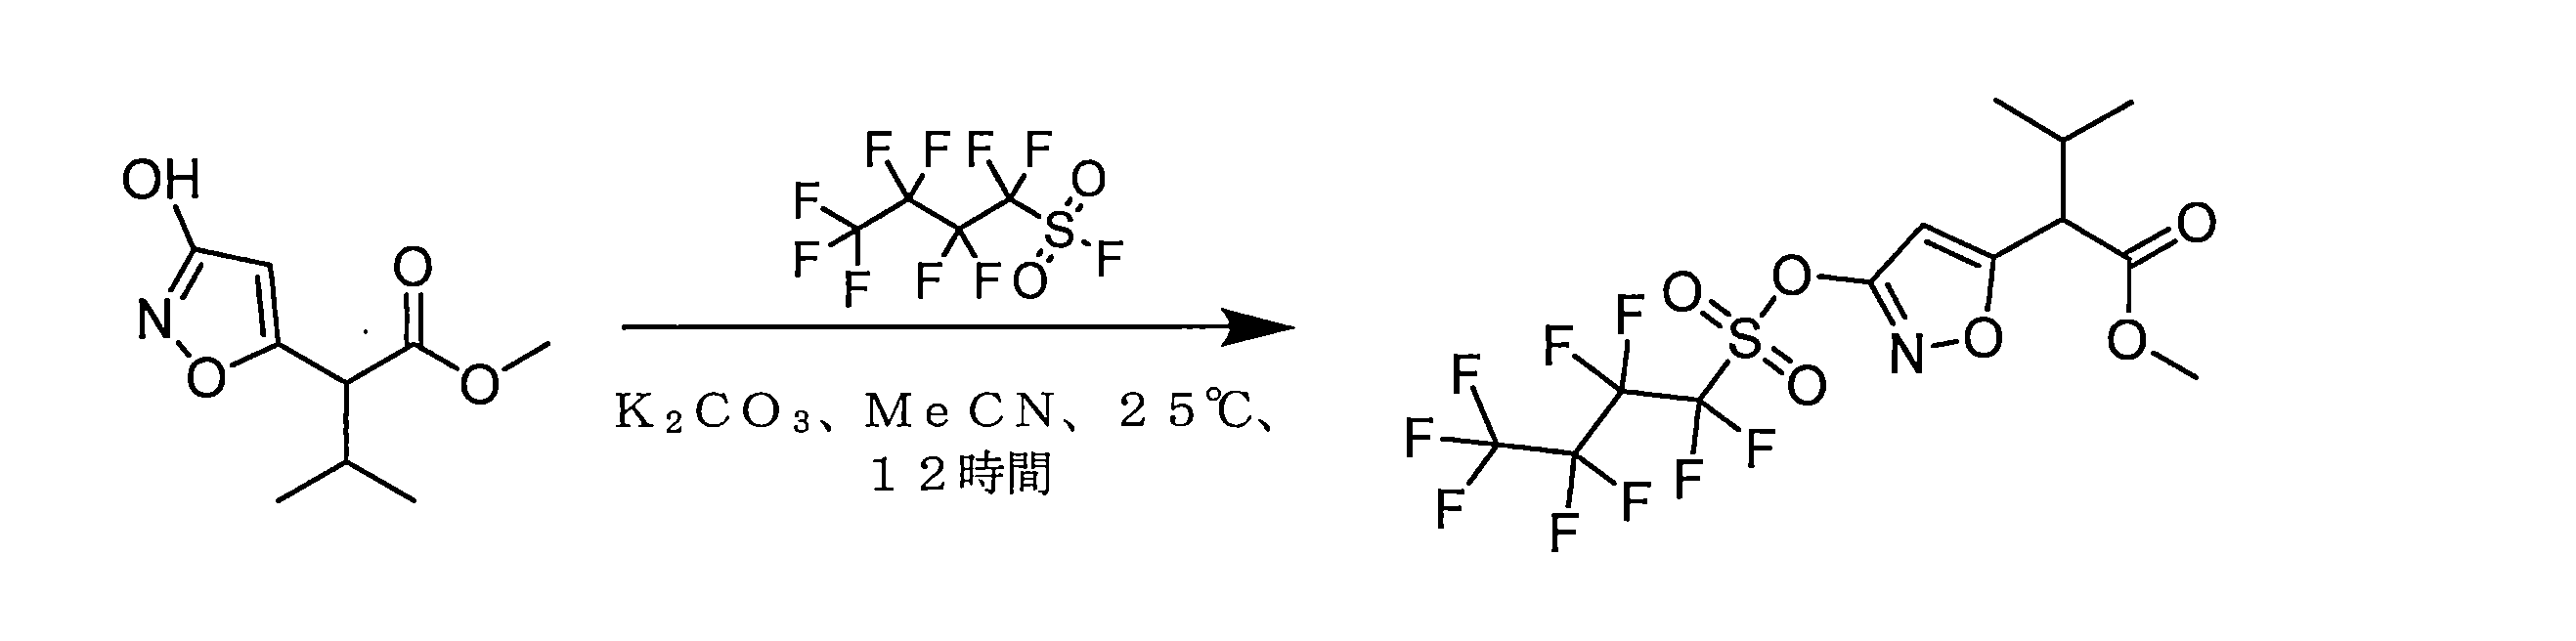
MeCN(1.6L)中の2-(3-メチルイソキサゾール-5-イル)-3-メチルブタン酸メチル(265g、1.33mol)[例示的な化合物144について記載されるように調製した〕の溶液に、K2CO3(367.71g、2.66mol)および1,1,2,2,3,3,4,4,4-ノナフルオロブタン-1-スルホニルフルオリド(482.25g、1.60mol、280.38mL)を添加した。反応混合物を、25℃で12時間撹拌した。反応混合物を減圧下で濃縮し、MeCNを除去した。残渣を水(1.5L)で希釈し、酢酸エチル(500mL×3)で抽出した。合わせた有機層を、ブライン(200mL)で洗浄し、Na2SO4上で乾燥させ、濾過し、減圧下で濃縮して、3-メチル-2-(3-(((ペルフルオロブチル)スルホニル)オキシ)イソキサゾール-5-イル)ブタン酸メチル(458g、886.42mmol)を茶色の油として得た。 To a solution of methyl 2-(3-methylisoxazol-5-yl)-3-methylbutanoate (265 g, 1.33 mol) [prepared as described for exemplary compound 144] in MeCN (1.6 L), K2CO3 (367.71 g, 2.66 mol) and 1,1,2,2,3,3,4,4,4-nonafluorobutane-1-sulfonyl fluoride (482.25 g, 1.60 mol, 280.38 mL) were added. The reaction mixture was stirred at 25 °C for 12 h. The reaction mixture was concentrated under reduced pressure to remove MeCN. The residue was diluted with water (1.5 L) and extracted with ethyl acetate (500 mL x 3). The combined organic layers were washed with brine (200 mL), dried over Na2SO4, filtered, and concentrated under reduced pressure to give methyl 3-methyl-2-(3-(((perfluorobutyl)sulfonyl)oxy)isoxazol-5-yl)butanoate (458 g, 886.42 mmol) as a brown oil.
工程2

DMF(1.6L)中の3-メチル-2-(3-(((ペルフルオロブチル)スルホニル)オキシ)イソキサゾール-5-イル)ブタン酸メチル(160g、332.44mmol)および4-(ジメトキシメチル)ピペリジン(79.40g、498.66mmol)の混合物に、DIEA(128.90g、997.32mmol、173.72mL)を、窒素下で25℃において一度に添加した。混合物を80℃で12時間撹拌した。混合物を25℃に冷却し、氷水(w:w=1:1)(3L)に注ぎ入れ、15分間撹拌した。水相を酢酸エチル(1L×5)で抽出した。合わせた有機相をブライン(300mL×3)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、真空中で濃縮した。混合物をEtOAc(70mL)中に溶解し、濾過して、不溶性材料を除去した。残渣を、カラムクロマトグラフィー(石油エーテル/酢酸エチル=80:1~0:1)によって精製して、(34g、100mmol)の2-(3-(4-(ジメトキシメチル)ピペリジン-1-イル)イソキサゾール-5-イル)-3-メチルブタン酸メチルをオレンジ色のシロップとして得た。 To a mixture of methyl 3-methyl-2-(3-(((perfluorobutyl)sulfonyl)oxy)isoxazol-5-yl)butanoate (160 g, 332.44 mmol) and 4-(dimethoxymethyl)piperidine (79.40 g, 498.66 mmol) in DMF (1.6 L) was added DIEA (128.90 g, 997.32 mmol, 173.72 mL) in one portion at 25° C. under nitrogen. The mixture was stirred at 80° C. for 12 h. The mixture was cooled to 25° C. and poured into ice water (w:w=1:1) (3 L) and stirred for 15 min. The aqueous phase was extracted with ethyl acetate (1 L×5). The combined organic phase was washed with brine (300 mL×3), dried over anhydrous sodium sulfate, filtered and concentrated in vacuo. The mixture was dissolved in EtOAc (70 mL) and filtered to remove insoluble material. The residue was purified by column chromatography (petroleum ether/ethyl acetate = 80:1 to 0:1) to obtain methyl 2-(3-(4-(dimethoxymethyl)piperidin-1-yl)isoxazol-5-yl)-3-methylbutanoate (34 g, 100 mmol) as an orange syrup.
工程3

MeOH(30mL)中の2-[3-[4-(ジメトキシメチル)-1-ピペリジル]イソキサゾール-5-イル]-3-メチル-ブタン酸メチル(10g、29.38mmol、1当量)の溶液に、水性LiOH溶液(6M、30.00mL、6.13当量)およびTHF(30mL)を添加した。混合物を25℃で1時間撹拌した。反応混合物のpHを1M HCl(200mL)によって調整し、水(200mL)で希釈し、EtOAc(200mL×3)で抽出した。合わせた有機層をブライン(200mL)で洗浄し、無水Na2SO4上で乾燥させ、濾過し、減圧下で濃縮した。粗2-[3-[4-(ジメトキシメチル)-1-ピペリジル]イソキサゾール-5-イル]-3-メチル-ブタン酸(9.5g、29.11mmol)を黄色の油として得た。 To a solution of methyl 2-[3-[4-(dimethoxymethyl)-1-piperidyl]isoxazol-5-yl]-3-methyl-butanoate (10 g, 29.38 mmol, 1 equiv.) in MeOH (30 mL), aqueous LiOH solution (6 M, 30.00 mL, 6.13 equiv.) and THF (30 mL) were added. The mixture was stirred at 25 °C for 1 h. The pH of the reaction mixture was adjusted by 1 M HCl (200 mL), diluted with water (200 mL), and extracted with EtOAc (200 mL x 3). The combined organic layers were washed with brine (200 mL), dried over anhydrous Na2SO4, filtered, and concentrated under reduced pressure. Crude 2-[3-[4-(dimethoxymethyl)-1-piperidyl]isoxazol-5-yl]-3-methyl-butanoic acid (9.5 g, 29.11 mmol) was obtained as a yellow oil.
工程4

DMF(100mL)中の(2S,4R)-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド塩酸塩(10.71g、29.11mmol、1当量)、DIPEA(11.29g、87.32mmol、15.21mL、3当量)、および2-[3-[4-(ジメトキシメチル)-1-ピペリジル]イソキサゾール-5-イル]-3-メチル-ブタン酸(9.5g、29.11mmol、1当量)の溶液に、HATU(16.60g、43.66mmol、1.5当量)を、0℃において窒素下で添加した。混合物を25℃に温め、1時間撹拌した。反応混合物を水(300mL)で希釈し、EtOAc(150mL×3)で抽出した。合わせた有機層を、ブライン(100mL)で洗浄し、Na2SO4上で乾燥させ、濾過し、減圧下で濃縮して、残渣を生成した。残渣を、分取HPLC(カラム:Phenomenex luna C18 250*50mm*10um;移動相:[水(0.225%FA)-MeCN];B%:30%~60%、15分)によって精製した。(2S,4R)-1-[2-[3-[4-(ジメトキシメチル)-1-ピペリジル]イソキサゾール-5-イル]-3-メチル-ブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(18g、27.57mmol)を白色の固体として得た。 To a solution of (2S,4R)-4-hydroxy-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide hydrochloride (10.71 g, 29.11 mmol, 1 eq.), DIPEA (11.29 g, 87.32 mmol, 15.21 mL, 3 eq.), and 2-[3-[4-(dimethoxymethyl)-1-piperidyl]isoxazol-5-yl]-3-methyl-butanoic acid (9.5 g, 29.11 mmol, 1 eq.) in DMF (100 mL), HATU (16.60 g, 43.66 mmol, 1.5 eq.) was added under nitrogen at 0°C. The mixture was warmed to 25°C and stirred for 1 h. The reaction mixture was diluted with water (300 mL) and extracted with EtOAc (150 mL x 3). The combined organic layers were washed with brine (100 mL), dried over Na2SO4, filtered and concentrated under reduced pressure to yield a residue. The residue was purified by preparative HPLC (column: Phenomenex luna C18 250*50 mm*10 um; mobile phase: [water (0.225% FA)-MeCN]; B%: 30%-60%, 15 min). (2S,4R)-1-[2-[3-[4-(dimethoxymethyl)-1-piperidyl]isoxazol-5-yl]-3-methyl-butanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (18 g, 27.57 mmol) was obtained as a white solid.
工程5
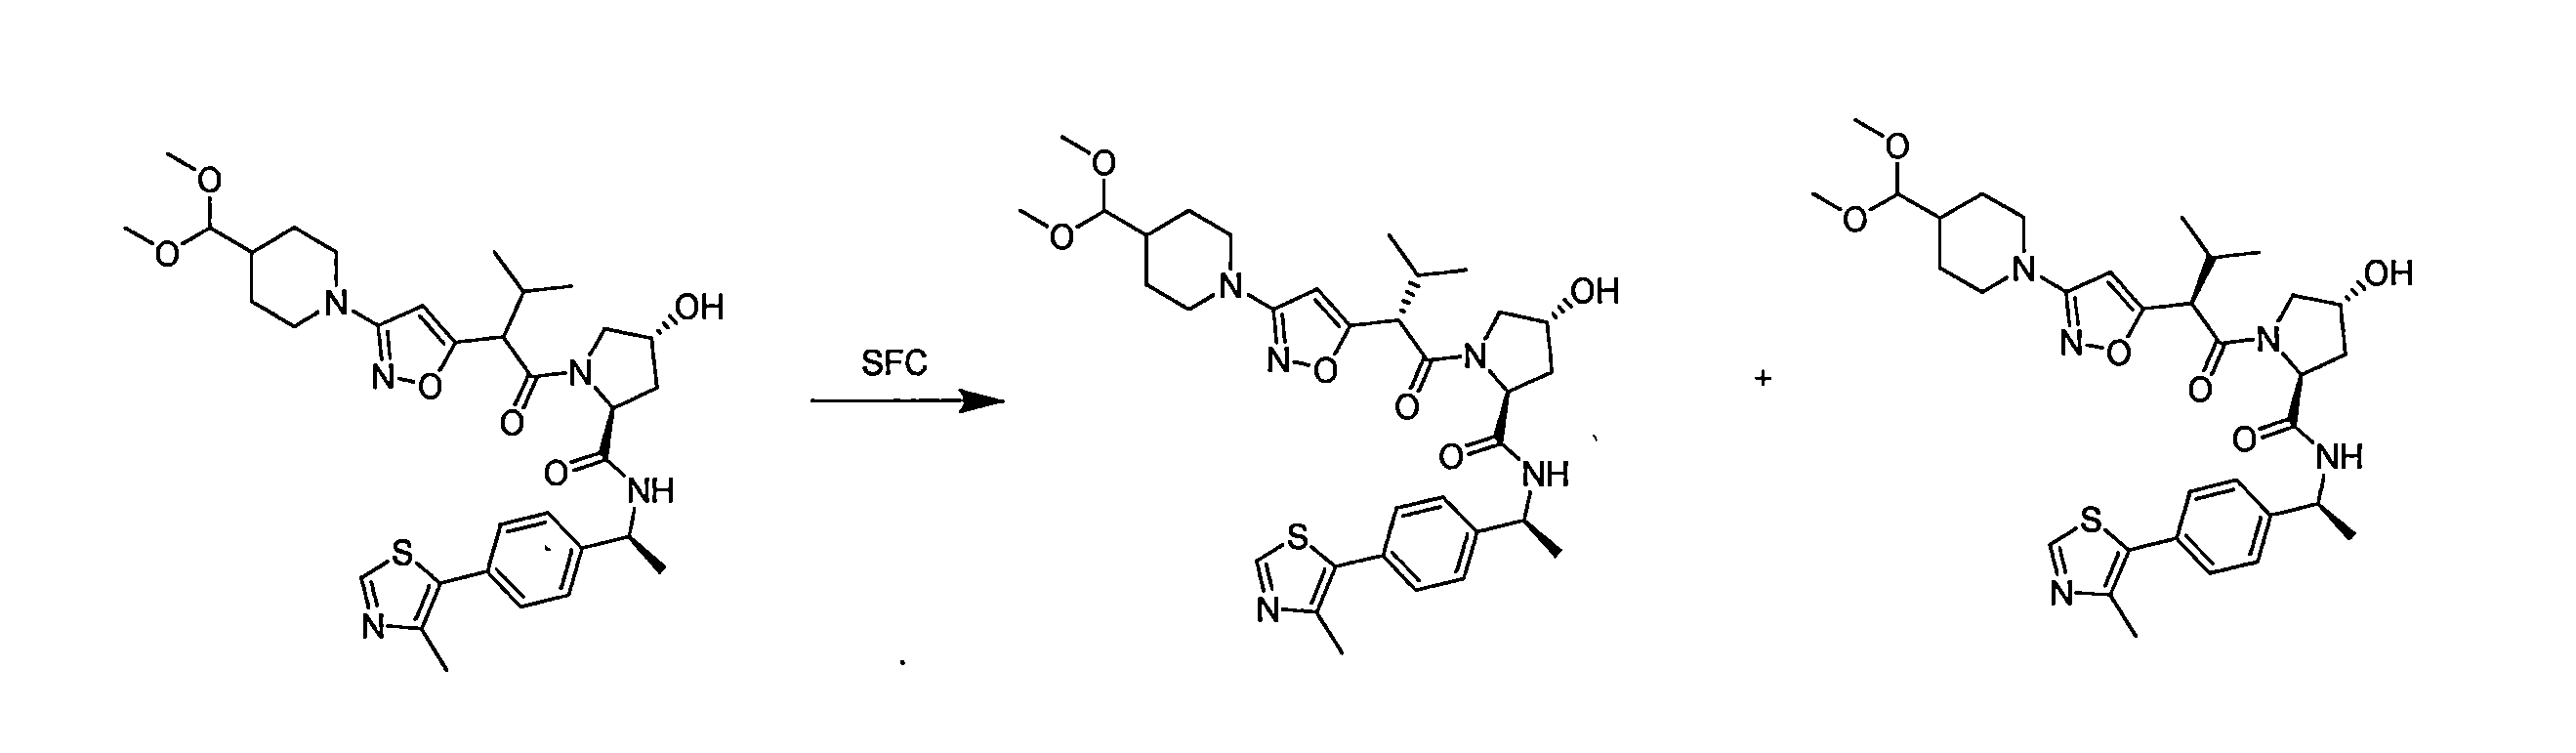
(2S,4R)-1-[2-[3-[4-(ジメトキシメチル)-1-ピペリジル]イソキサゾール-5-イル]-3-メチル-ブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(5g、7.81mmol、1当量)を、分取SFC(カラム:DAICEL CHIRALPAK AD-H 250mm×30mm、I.D.、5um;移動相:CO2中のイソプロパノール(0.1%NH3H2O)40%~40%;流速:70g/分;波長:220nm)および(カラム:DAICEL CHIRALPAK AD 250mm×30mm、I.D.、10um;移動相:CO2中のイソプロパノール(0.1%NH3H2O)40%~40%;流速:70g/分;波長:220nm)によって2回分離した。 (2S,4R)-1-[2-[3-[4-(dimethoxymethyl)-1-piperidyl]isoxazol-5-yl]-3-methyl-butanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (5 g, 7.81 mmol, 1 eq.) was purified by preparative SFC (column: DAICEL CHIRALPAK AD-H 250 mm x 30 mm, I.D., 5 um; mobile phase: isopropanol (0.1% NH3H2O) 40%-40% in CO2; flow rate: 70 g/min; wavelength: 220 nm) and (column: DAICEL CHIRALPAK AD 250 mm x 30 mm, I.D., 10 um; mobile phase: isopropanol (0.1% NH3H2O) 40%-40% in CO2; flow rate: 70 g/min; wavelength: 220 nm) and separated twice.
化合物(2S,4R)-1-[(2S)-2-[3-[4-(ジメトキシメチル)-1-ピペリジル]イソキサゾール-5-イル]-3-メチル-ブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(2.5g、3.91mmol)を白色の固体として得た。 The compound (2S,4R)-1-[(2S)-2-[3-[4-(dimethoxymethyl)-1-piperidyl]isoxazol-5-yl]-3-methyl-butanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (2.5 g, 3.91 mmol) was obtained as a white solid.
化合物(2S,4R)-1-[(2R)-2-[3-[4-(ジメトキシメチル)-1-ピペリジル]イソキサゾール-5-イル]-3-メチル-ブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(2.4g、3.68mmol)を白色の固体として得た。 The compound (2S,4R)-1-[(2R)-2-[3-[4-(dimethoxymethyl)-1-piperidyl]isoxazol-5-yl]-3-methyl-butanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (2.4 g, 3.68 mmol) was obtained as a white solid.
工程6
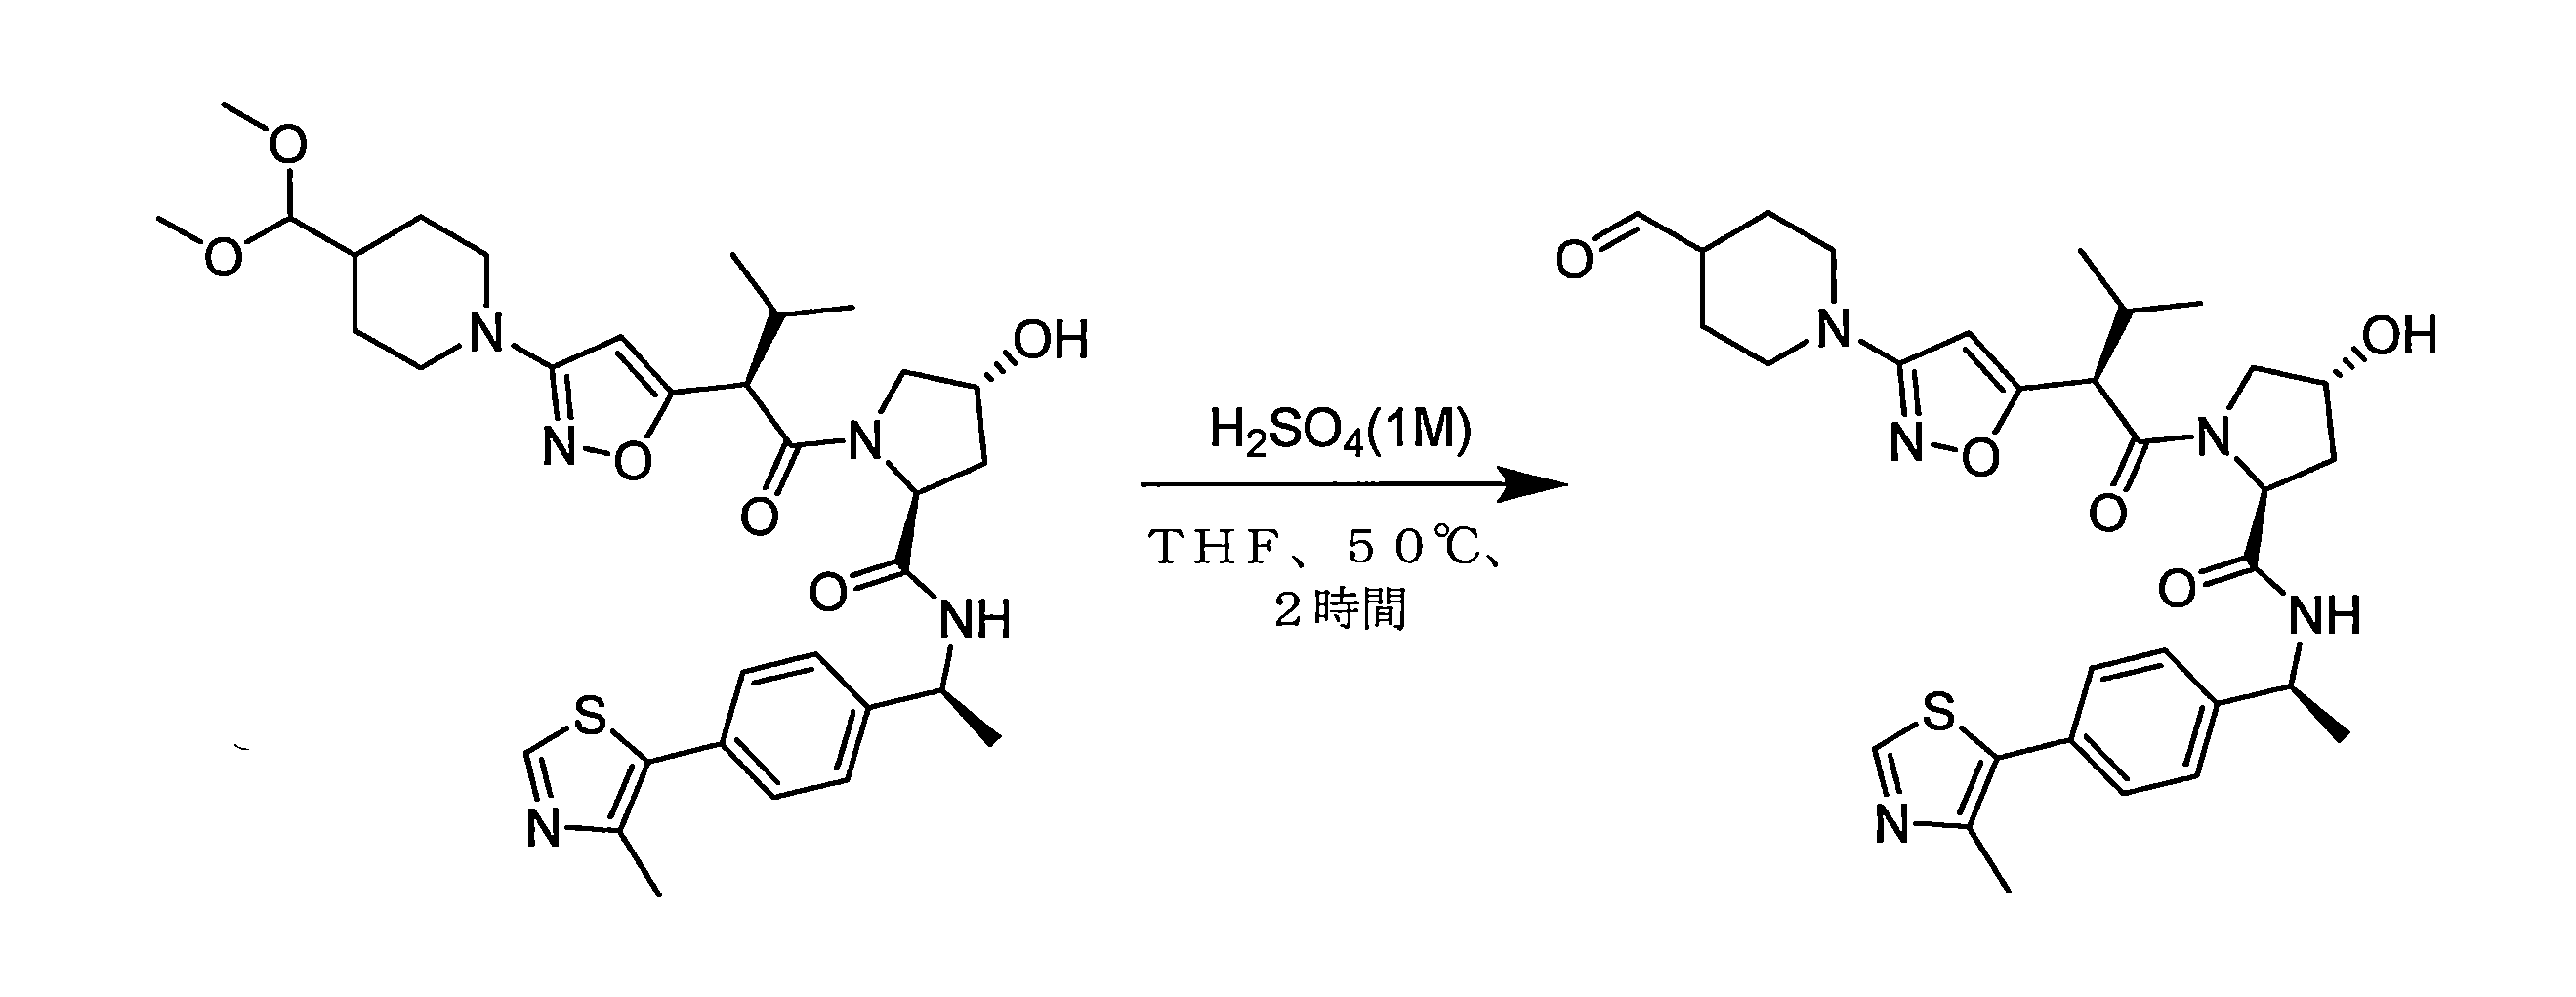
THF(15mL)中の(2S,4R)-1-[(2R)-2-[3-[4-ジメトキシメチル)-1-ピペリジル]イソキサゾール-5-イル]-3-メチル-ブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(1.2g、1.88mmol、1当量)の溶液に、H2SO4(1M、13.85mL、7.38当量)を添加した。混合物を50℃で2時間撹拌した。反応混合物を、飽和NaHCO3(30mL)を添加することによってクエンチし、次いで、水(30mL)で希釈し、ブライン(50mL×3)で抽出した。合わせた有機層を、ブライン(30mL)で洗浄し、無水Na2SO4上で乾燥させ、濾過し、減圧下で濃縮して、残渣を得た。粗(2S,4R)-1-[(2R)-2-[3-(4-ホルミル-1-ピペリジル)イソキサゾール-5-イル]-3-メチル-ブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(1.05g、1.77mmol)を白色の固体として得た。 To a solution of (2S,4R)-1-[(2R)-2-[3-[4-dimethoxymethyl)-1-piperidyl]isoxazol-5-yl]-3-methyl-butanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (1.2 g, 1.88 mmol, 1 equiv.) in THF (15 mL) was added H2SO4 (1 M, 13.85 mL, 7.38 equiv.). The mixture was stirred at 50°C for 2 h. The reaction mixture was quenched by adding saturated NaHCO3 (30 mL), then diluted with water (30 mL) and extracted with brine (50 mL x 3). The combined organic layers were washed with brine (30 mL), dried over anhydrous Na2SO4, filtered and concentrated under reduced pressure to give a residue. Crude (2S,4R)-1-[(2R)-2-[3-(4-formyl-1-piperidyl)isoxazol-5-yl]-3-methyl-butanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (1.05 g, 1.77 mmol) was obtained as a white solid.
工程7

メタノール(40mL)中の2-[6-アミノ-5-[8-[2-[3-(4-ピペリジルオキシ)シクロブトキシ]-4-ピリジル]-3,8-ジアザビシクロ[3.2.1]オクタン-3-イル]ピリダジン-3-イル]フェノール二塩酸塩(2.08g、3.37mmol、1当量)の溶液に、酢酸ナトリウム(829mg、10.11mmol、3当量)を添加し、次いで、混合物に、(2S,4R)-1-[(2R)-2-[3-(4-ホルミル-1-ピペリジル)イソキサゾール-5-イル]-3-メチル-ブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミド(2g、3.37mmol、1当量)および酢酸(23mg、0.33mmol、0.1当量)を添加した。混合物を25℃で0.5時間撹拌し、次いで、混合物に、シアノホウ化水素ナトリウム(1.06g、16.84mmol、5当量)を添加した。混合物を25℃で2時間撹拌した。反応混合物を減圧下で濃縮した。残渣を、分取HPLC(カラム:Phenomenex luna C18 250*50mm*10um;移動相:[水(0.225%FA)-ACN];B%:10%~40%、22分)によって精製した。(2S,4R)-1-[(2R)-2-[3-[4-[[4-[3-[[4-[3-[3-アミノ-6-(2-ヒドロキシフェニル)ピリダジン-4-イル]-3,8-ジアザビシクロ[3.2.1]オクタン-8-イル]-2-ピリジル]オキシ]シクロブトキシ]-1-ピペリジル]メチル]-1-ピペリジル]イソキサゾール-5-イル]-3-メチル-ブタノイル]-4-ヒドロキシ-N-[(1S)-1-[4-(4-メチルチアゾール-5-イル)フェニル]エチル]ピロリジン-2-カルボキサミドギ酸塩(1196mg、1.02mmol)を白色の固体として得た。 To a solution of 2-[6-amino-5-[8-[2-[3-(4-piperidyloxy)cyclobutoxy]-4-pyridyl]-3,8-diazabicyclo[3.2.1]octan-3-yl]pyridazin-3-yl]phenol dihydrochloride (2.08 g, 3.37 mmol, 1 eq.) in methanol (40 mL), sodium acetate (829 mg, 10.11 mmol, 3 eq.) was added, and the mixture was then cooled to 100° C. To the mixture was added (2S,4R)-1-[(2R)-2-[3-(4-formyl-1-piperidyl)isoxazol-5-yl]-3-methyl-butanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide (2 g, 3.37 mmol, 1 eq.) and acetic acid (23 mg, 0.33 mmol, 0.1 eq.). The mixture was stirred at 25° C. for 0.5 h, and then to the mixture was added sodium cyanoborohydride (1.06 g, 16.84 mmol, 5 eq.). The mixture was stirred at 25° C. for 2 h. The reaction mixture was concentrated under reduced pressure. The residue was purified by preparative HPLC (column: Phenomenex luna C18 250*50mm*10um; mobile phase: [water (0.225% FA)-ACN]; B%: 10%-40%, 22 min). (2S,4R)-1-[(2R)-2-[3-[4-[[4-[3-[[4-[3-[3-amino-6-(2-hydroxyphenyl)pyridazin-4-yl]-3,8-diazabicyclo[3.2.1]octan-8-yl]-2-pyridyl]oxy]cyclobutoxy]-1-piperidyl]methyl]-1-piperidyl]isoxazol-5-yl]-3-methyl-butanoyl]-4-hydroxy-N-[(1S)-1-[4-(4-methylthiazol-5-yl)phenyl]ethyl]pyrrolidine-2-carboxamide formate (1196 mg, 1.02 mmol) was obtained as a white solid.
例示的な化合物594の例示的な合成
上記の他の実施例について記載されたものと類似の手順を使用して、以下のスキームに従って調製した。

例示的な化合物596は、類似の手順を使用して調製され得る。 Exemplary compound 596 can be prepared using a similar procedure.
例示的な化合物602の例示的な合成
例示的な化合物143について記載されるものと類似の手順を使用して、以下のスキームに従って調製した。

例示的な化合物607の例示的な合成
例示的な化合物143および例示的な化合物178について記載される手順を使用して調製した。
Exemplary Synthesis of Exemplary Compound 607 Prepared using procedures described for Exemplary Compound 143 and Exemplary Compound 178.
例示的な化合物610の例示的な合成
例示的な化合物143および例示的な化合物188、ならびに上記の他の実施例について記載されたものと類似の手順を使用して、以下のスキームに従って調製した。
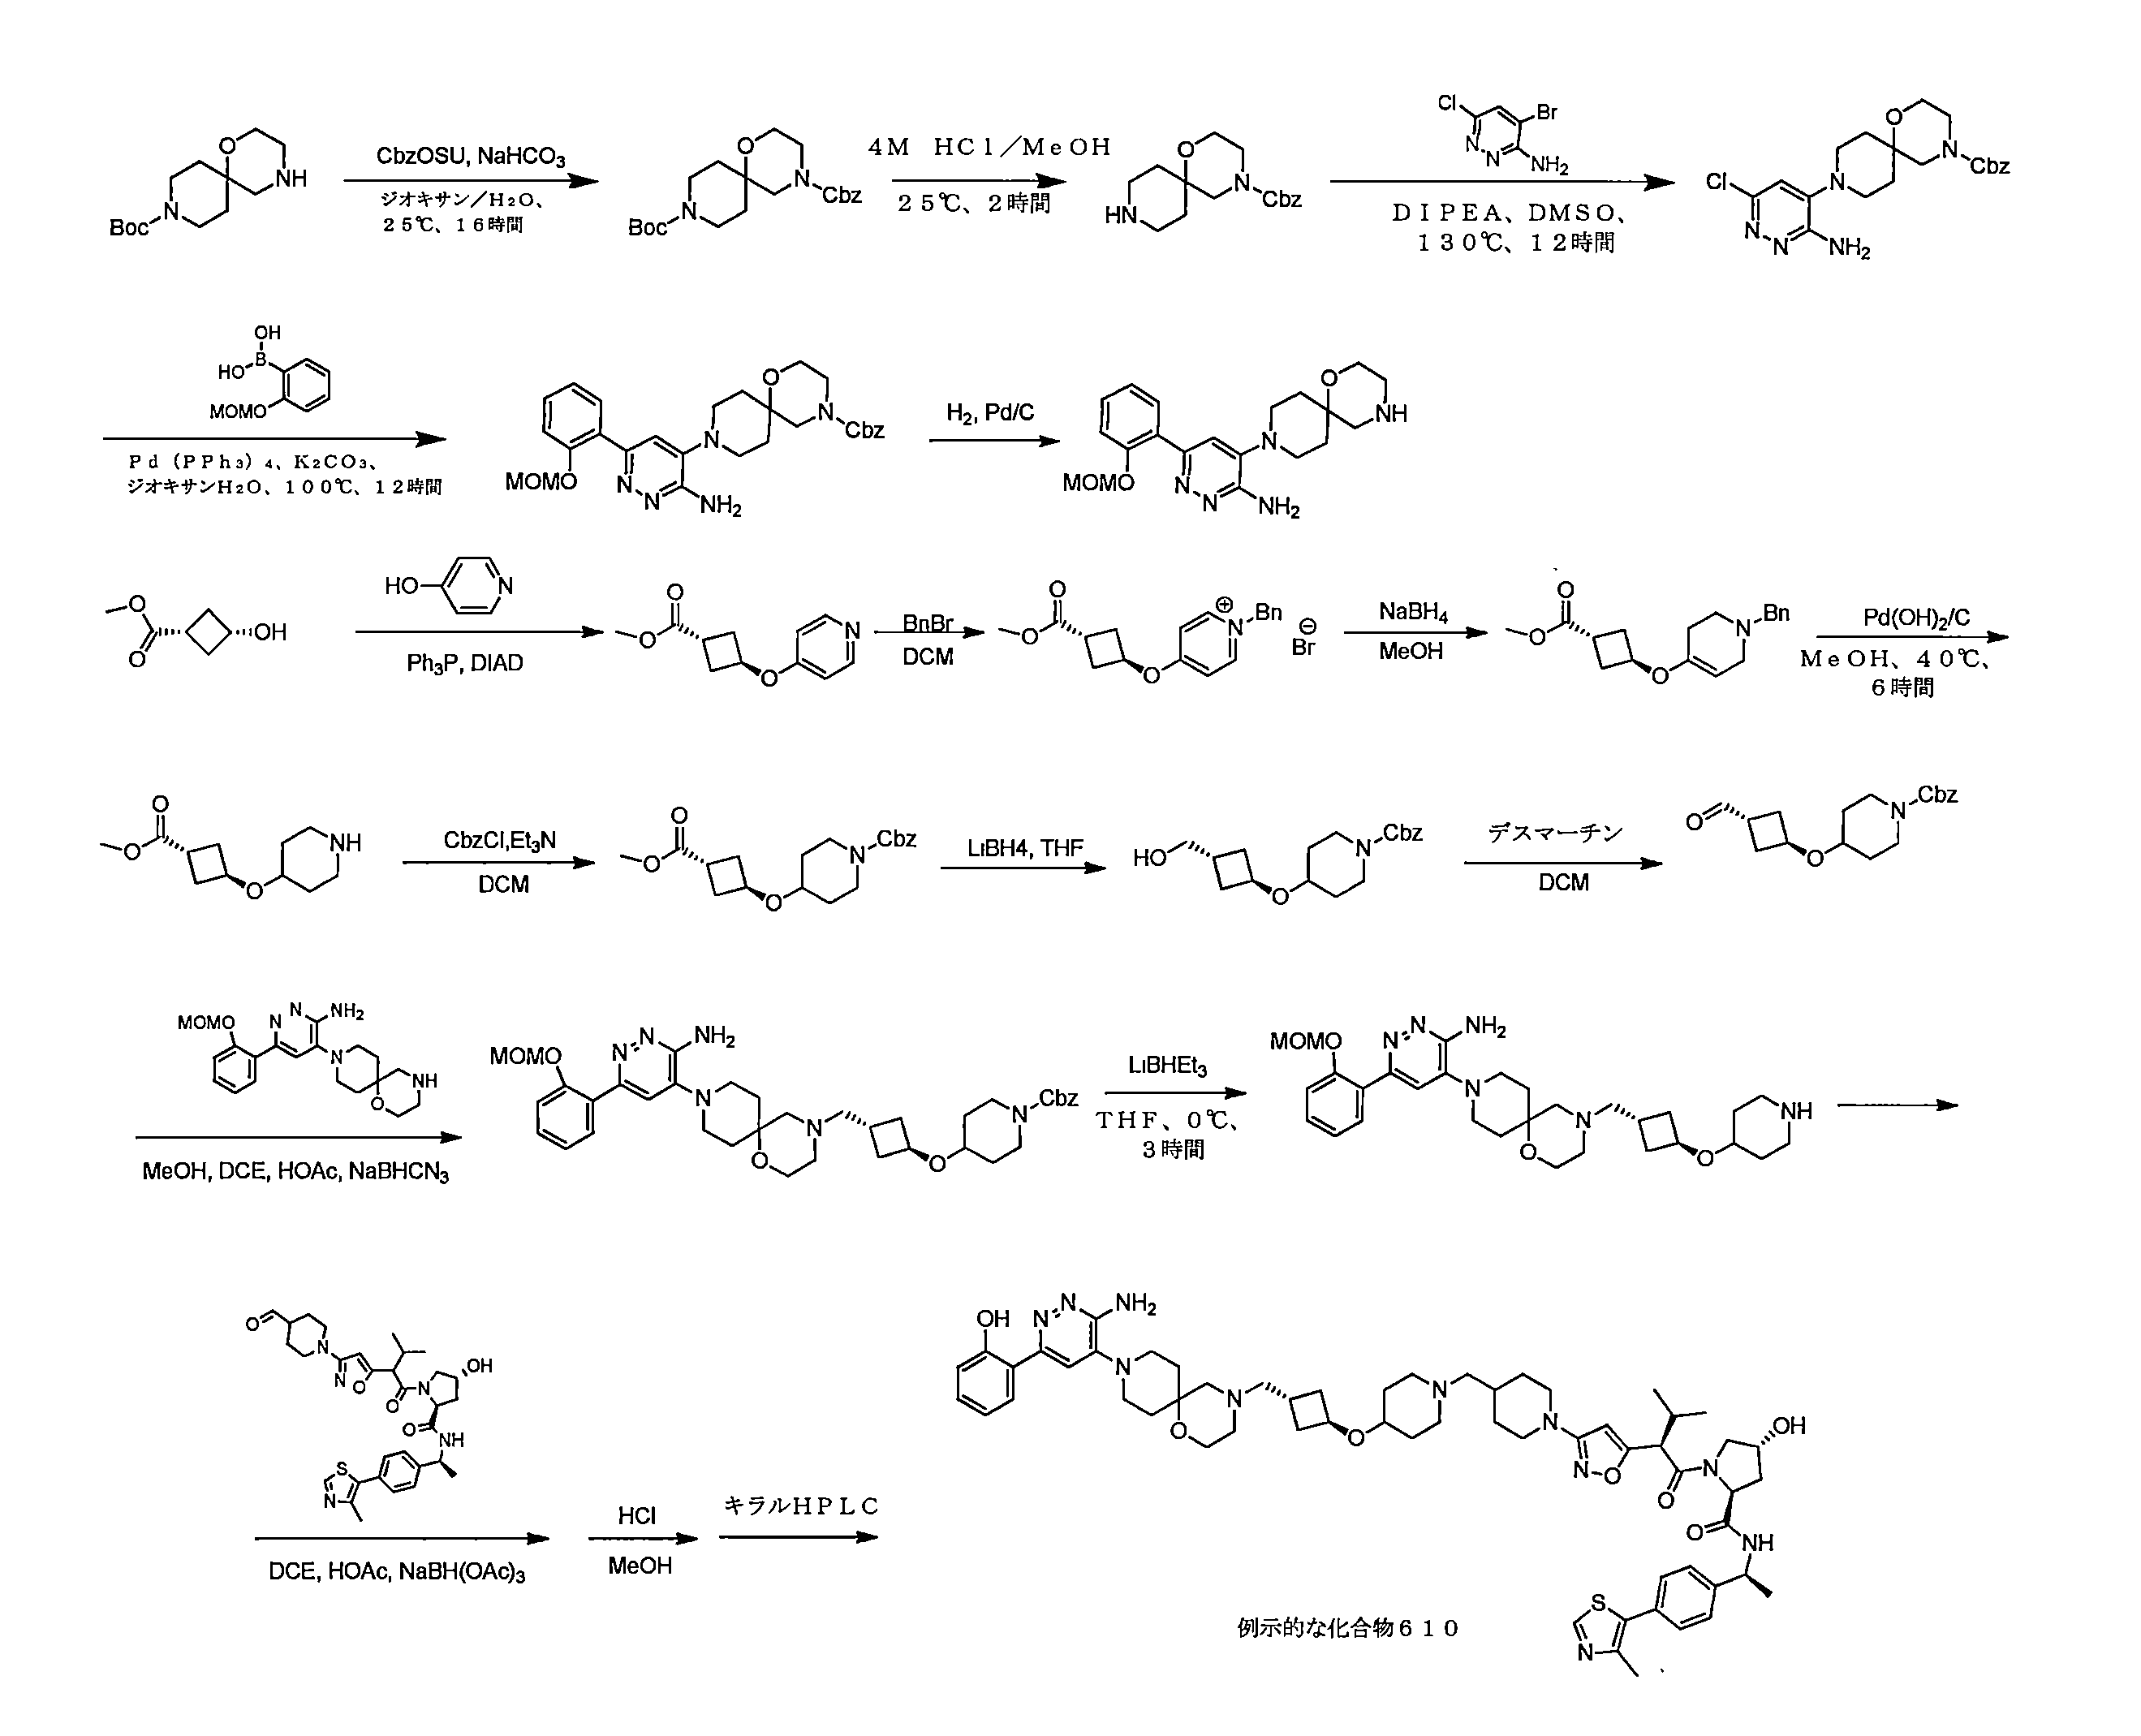
例示的な化合物616の例示的な合成
上記の実施例について記載されたものと類似の手順を使用して、以下のスキームに従って調製した。
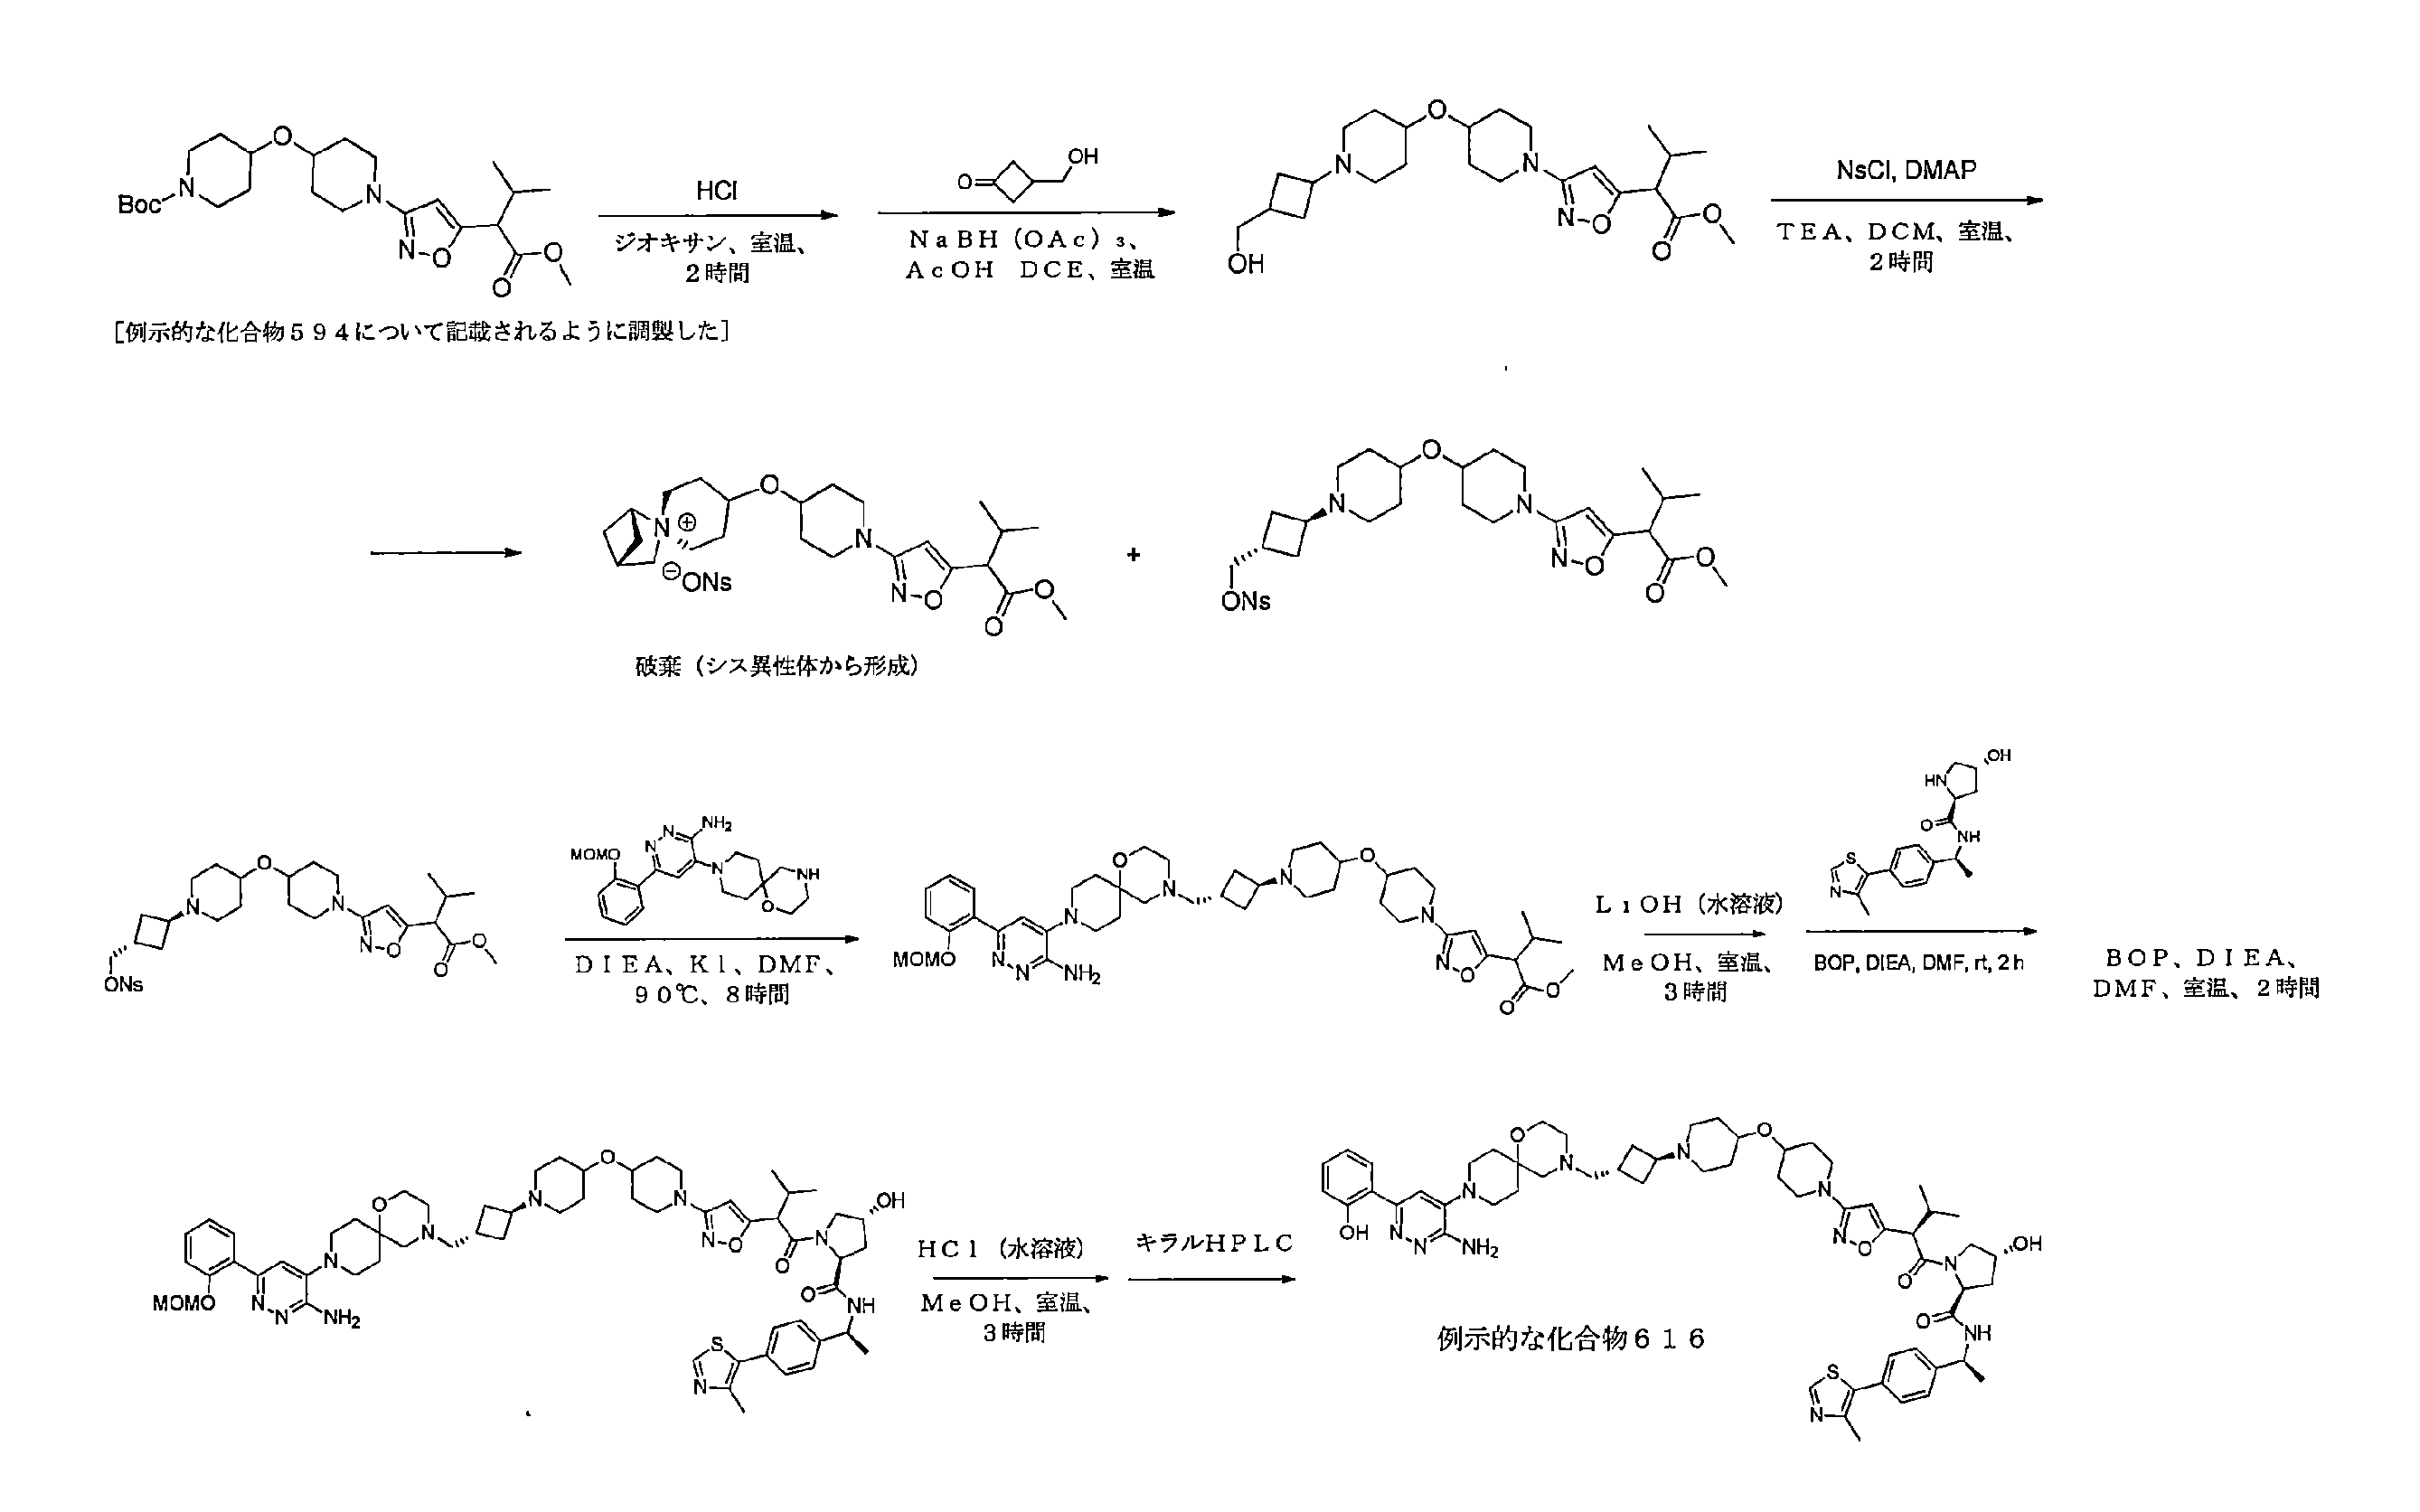
例示的な化合物618の例示的な合成
例示的な化合物610および例示的な化合物143、ならびに上記の他の実施例について記載された手順を使用して、以下のスキームに従って調製した。
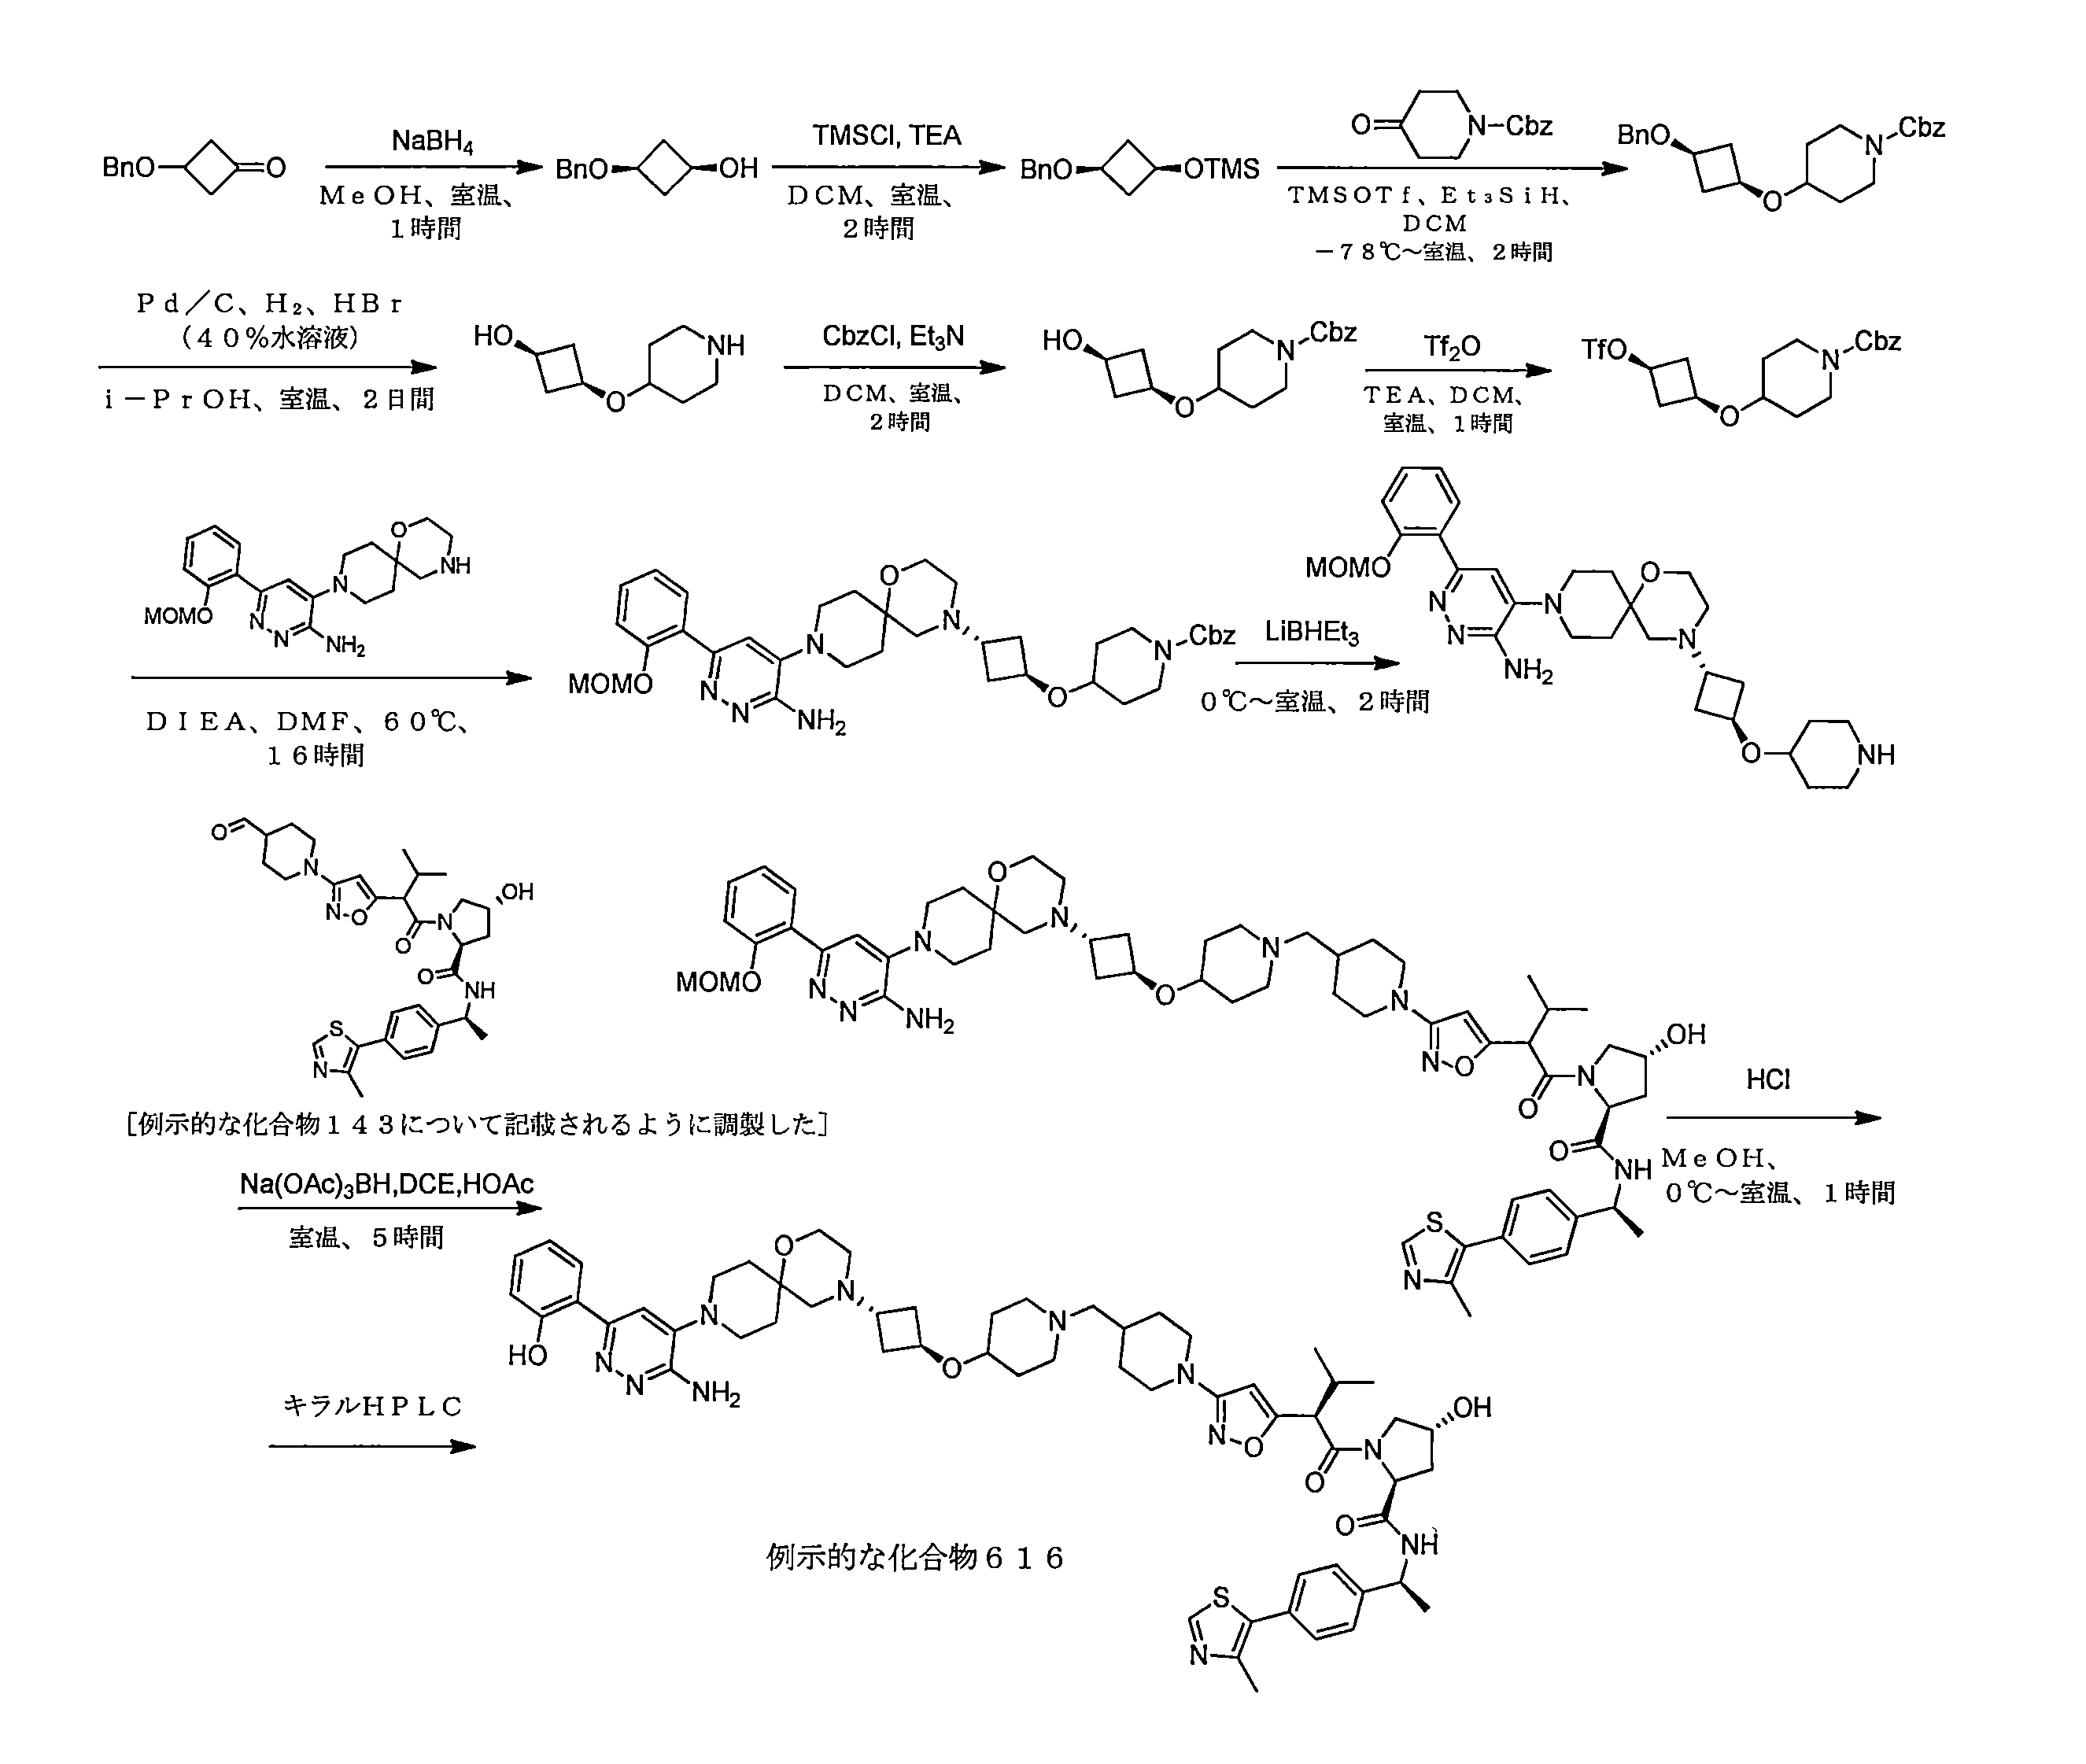
例示的な化合物626の例示的な合成
工程1

窒素不活性雰囲気下でパージし、維持した250mLの丸底フラスコに、tert-ブチル4-[(1r,3r)-3-ヒドロキシシクロブトキシ]ピペリジン-1-カルボキシレート(5g、18.43mmol、1.0当量)、DMF(100mL)を入れ、NaH(2.22g、55.42mmol、3.0当量、油中60%)を0℃で添加した。これを室温で15分間反応させた。これに続いて、4-ブロモ-2-フルオロピリジン(3.57g、20.27mmol、1.1当量)を添加した。得られた溶液を室温で4時間撹拌した。次いで、100mLの水/氷を添加することによって反応をクエンチさせた。得られた溶液を(100mL×3)の酢酸エチルで抽出し、有機層を合わせた。得られた混合物を(100mL×1)のブラインで洗浄した。混合物を、無水硫酸ナトリウムで乾燥させた。残渣を、酢酸エチル/石油エーテル(1/2)で溶出するシリカゲルカラム上に適用した。収集した画分をまとめ、真空下で濃縮した。これにより、6.3g(80%)のtert-ブチル4-[(1r,3r)-3-[(4-ブロモピリジン-2-イル)オキシ]シクロブトキシ]ピペリジン-1-カルボキシレートを黄色の油として得た。 A 250 mL round bottom flask purged and maintained under nitrogen inert atmosphere was charged with tert-butyl 4-[(1r,3r)-3-hydroxycyclobutoxy]piperidine-1-carboxylate (5 g, 18.43 mmol, 1.0 equiv), DMF (100 mL) and NaH (2.22 g, 55.42 mmol, 3.0 equiv, 60% in oil) was added at 0°C. This was allowed to react at room temperature for 15 minutes. This was followed by the addition of 4-bromo-2-fluoropyridine (3.57 g, 20.27 mmol, 1.1 equiv). The resulting solution was stirred at room temperature for 4 hours. The reaction was then quenched by adding 100 mL of water/ice. The resulting solution was extracted with (100 mL x 3) ethyl acetate and the organic layers were combined. The resulting mixture was washed with (100 mL x 1) brine. The mixture was dried over anhydrous sodium sulfate. The residue was applied onto a silica gel column eluted with ethyl acetate/petroleum ether (1/2). The collected fractions were combined and concentrated under vacuum. This gave 6.3 g (80%) of tert-butyl 4-[(1r,3r)-3-[(4-bromopyridin-2-yl)oxy]cyclobutoxy]piperidine-1-carboxylate as a yellow oil.
工程2
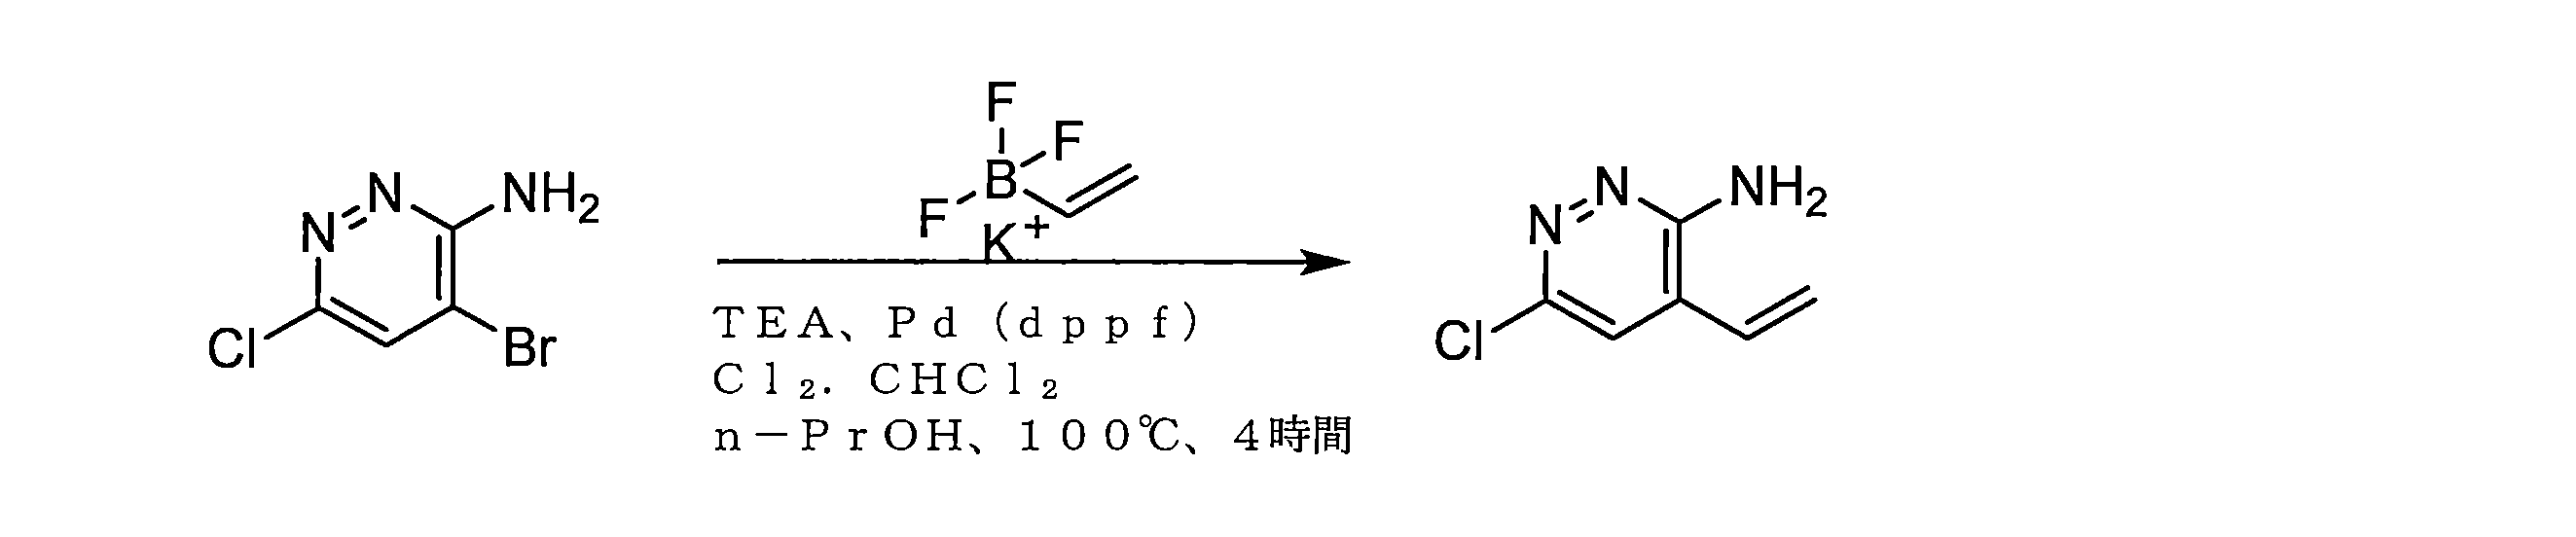
窒素不活性雰囲気下でパージし、維持した250mLの丸底フラスコに、4-ブロモ-6-クロロピリダジン-3-アミン(5g、24.0mmol、1.0当量)、n-プロパノール(100mL)、TEA(7.28g、72.0mmol、3当量)、トリフルオロビニルホウ酸カリウム(2.39g、25.0mmol、1.05当量)、およびPd(dppf)Cl2.CH2Cl2(1.96g、2.39mmol、0.10当量)を入れた。得られた溶液を油浴中、100℃で4時間撹拌した。固体を濾別した。残渣を、酢酸エチル/石油エーテル(1/1)とともにシリカゲルカラムに充填させた。収集した画分をまとめ、真空下で濃縮した。これにより、2.3g(62%)の6-クロロ-4-エテニルピリダジン-3-アミンを黄色の固体として得た。 A 250 mL round bottom flask purged and maintained under nitrogen inert atmosphere was charged with 4-bromo-6-chloropyridazin-3-amine (5 g, 24.0 mmol, 1.0 equiv), n-propanol (100 mL), TEA (7.28 g, 72.0 mmol, 3 equiv), potassium trifluorovinylborate (2.39 g, 25.0 mmol, 1.05 equiv), and Pd(dppf)Cl2.CH2Cl2 (1.96 g, 2.39 mmol, 0.10 equiv). The resulting solution was stirred in an oil bath at 100 °C for 4 h. The solids were filtered off. The residue was loaded onto a silica gel column with ethyl acetate/petroleum ether (1/1). The collected fractions were combined and concentrated under vacuum. This gave 2.3 g (62%) of 6-chloro-4-ethenylpyridazin-3-amine as a yellow solid.
工程3

窒素不活性雰囲気下でパージし、維持した20mLの密封チューブに、6-クロロ-4-エテニルピリダジン-3-アミン(910mg、5.84mmol、1.0当量)、DMF(14mL)、TEA(1775mg、17.5mmol、3.0当量)、tert-ブチル4-[(1r,3r)-3-[(4-ブロモピリジン-2-イル)オキシ]シクロブトキシ]ピペリジン-1-カルボキシレート(2.50g、5.849mmol、1.0当量)、トリ-o-トリルホスフィン(1780mg、5.85mmol、1.0当量)、Pd(OAc)2(131mg、0.585mmol、0.10当量)を入れた。得られた溶液を、油浴中で、130℃で16時間撹拌した。次いで、反応を30mLの水の添加によりクエンチさせた。得られた溶液を(20mL×3)の酢酸エチルで抽出し、有機層を合わせた。得られた混合物を、(20mL×3)のブラインで洗浄した。混合物を、無水硫酸ナトリウムで乾燥させた。残渣を、ジクロロメタン/メタノール(10/1)で溶出するシリカゲルカラム上に適用した。収集した画分をまとめ、真空下で濃縮した。これにより、625mg(21%)のtert-ブチル4-[(1r,3r)-3-([4-[(E)-2-(3-アミノ-6-クロロピリダジン-4-イル)エテニル]ピリジン-2-イル]オキシ)シクロブトキシ]ピペリジン-1-カルボキシレートを茶色の固体として得た。 In a 20 mL sealed tube purged and maintained under a nitrogen inert atmosphere was placed 6-chloro-4-ethenylpyridazin-3-amine (910 mg, 5.84 mmol, 1.0 equiv.), DMF (14 mL), TEA (1775 mg, 17.5 mmol, 3.0 equiv.), tert-butyl 4-[(1r,3r)-3-[(4-bromopyridin-2-yl)oxy]cyclobutoxy]piperidine-1-carboxylate (2.50 g, 5.849 mmol, 1.0 equiv.), tri-o-tolylphosphine (1780 mg, 5.85 mmol, 1.0 equiv.), Pd(OAc)2 (131 mg, 0.585 mmol, 0.10 equiv.). The resulting solution was stirred in an oil bath at 130 °C for 16 h. The reaction was then quenched by the addition of 30 mL of water. The resulting solution was extracted with ethyl acetate (20 mL x 3) and the organic layers were combined. The resulting mixture was washed with brine (20 mL x 3). The mixture was dried over anhydrous sodium sulfate. The residue was applied onto a silica gel column eluted with dichloromethane/methanol (10/1). The collected fractions were combined and concentrated under vacuum. This gave 625 mg (21%) of tert-butyl 4-[(1r,3r)-3-([4-[(E)-2-(3-amino-6-chloropyridazin-4-yl)ethenyl]pyridin-2-yl]oxy)cyclobutoxy]piperidine-1-carboxylate as a brown solid.
tert-ブチル4-[(1r,3r)-3-([4-[(E)-2-(3-アミノ-6-クロロピリダジン-4-イル)エテニル]ピリジン-2-イル]オキシ)シクロブトキシ]ピペリジン-1-カルボキシレートを、例示的な化合物143および上記の他の実施例について記載された手順を使用して、最終化合物に変換した。
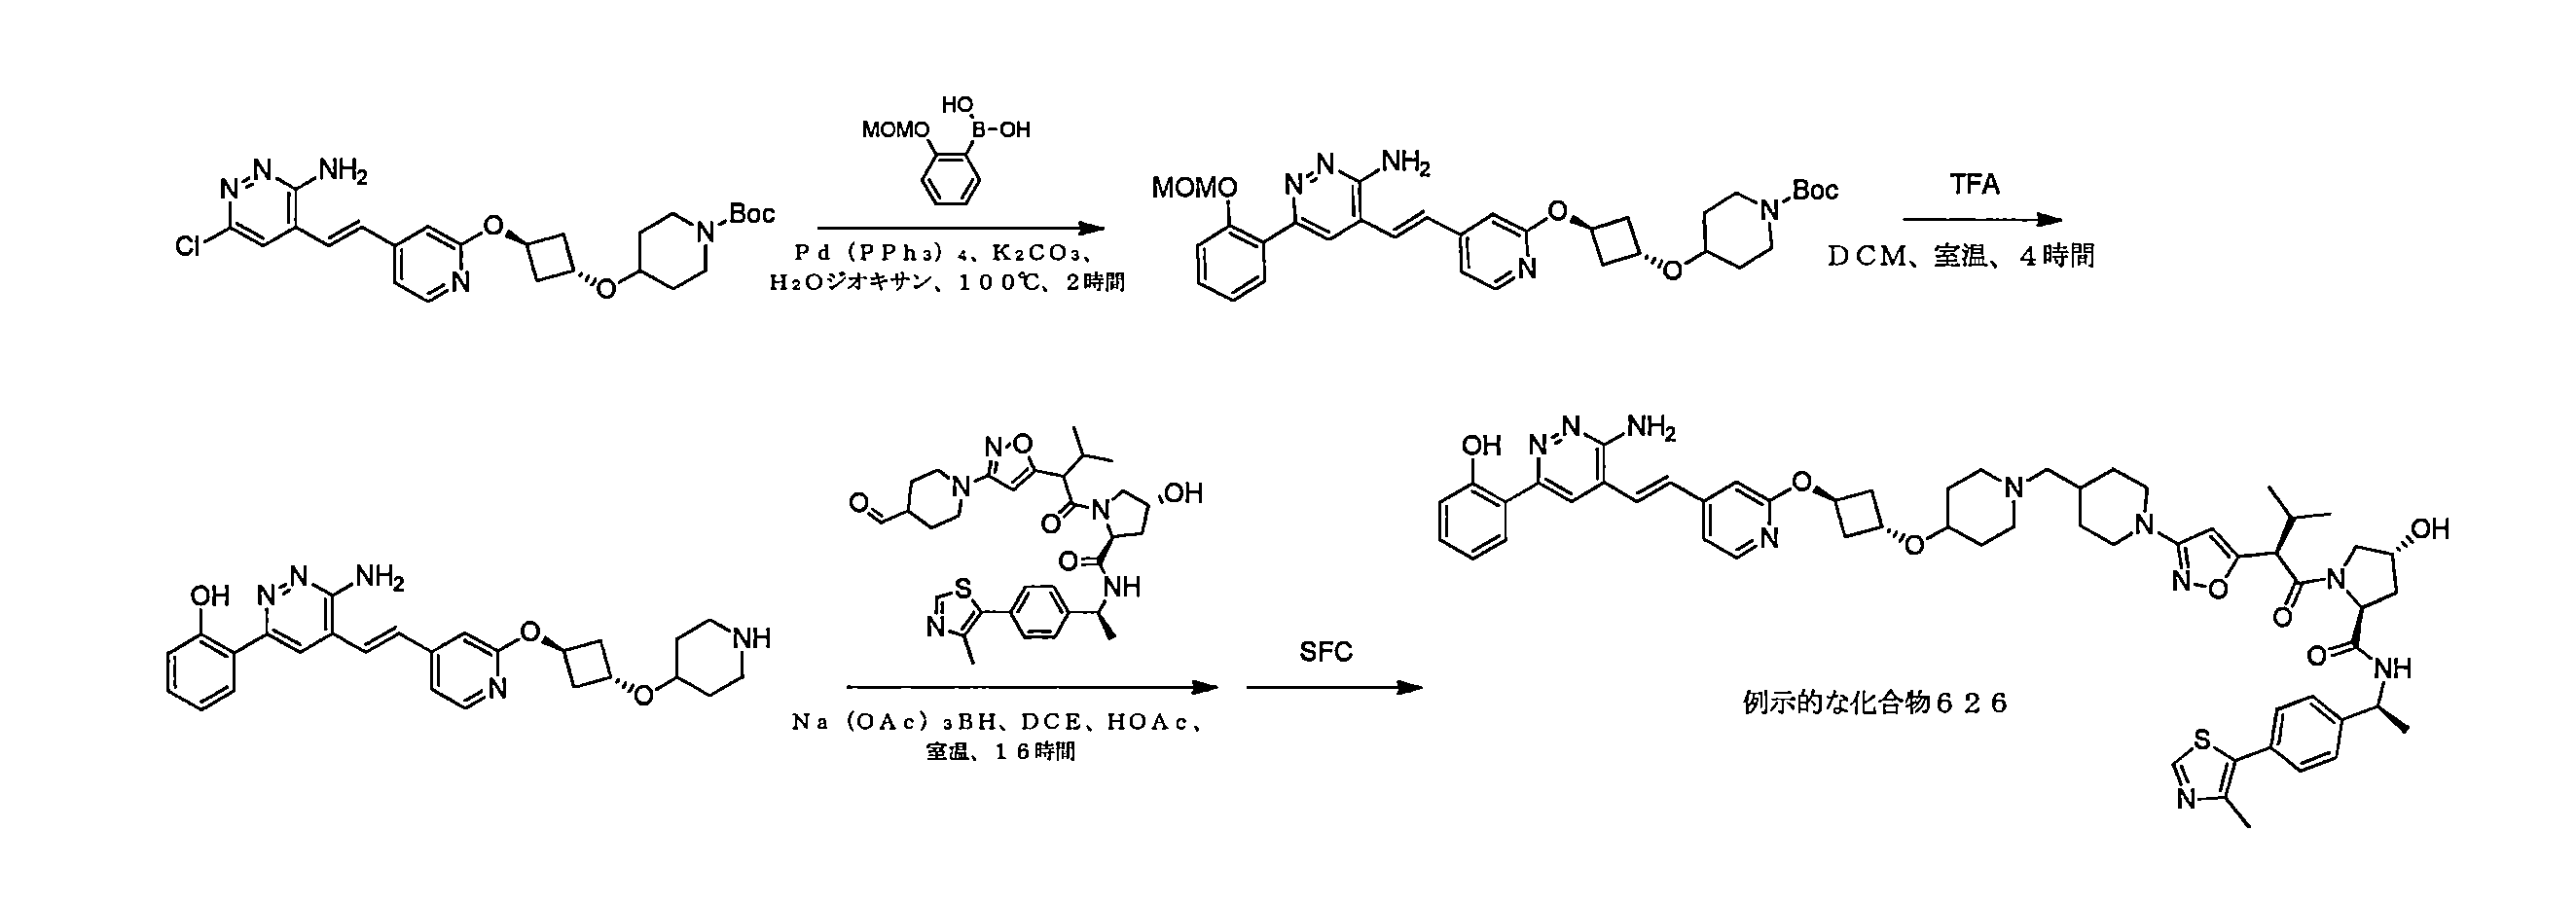
例示的な化合物630の例示的な合成
工程1

ジクロロメタン(30mL)中のメチル4-アミノチアゾール-5-カルボキシレート(0.4g、2.53mmol、1当量)およびトリエチルアミン(512mg、5.06mmol、0.7mL、2当量)の溶液に、ジ-tert-ブチルジカーボネート(1.10g、5.06mmol、1.2mL、2当量)および4-ジメチルアミノピリジン(62mg、0.50mmol、0.2当量)を窒素下で、25℃において一度に添加した。混合物を25℃で16時間撹拌した。混合物を、45℃において減圧下で濃縮した。残渣を、シリカゲルカラムクロマトグラフィー(石油エーテル/酢酸エチル=20/1~8/1)によって精製して、メチル4-[ビス(tert-ブトキシカルボニル)アミノ]チアゾール-5-カルボキシレート(850mg)を黄色の油として得た。 To a solution of methyl 4-aminothiazole-5-carboxylate (0.4 g, 2.53 mmol, 1 eq.) and triethylamine (512 mg, 5.06 mmol, 0.7 mL, 2 eq.) in dichloromethane (30 mL), di-tert-butyl dicarbonate (1.10 g, 5.06 mmol, 1.2 mL, 2 eq.) and 4-dimethylaminopyridine (62 mg, 0.50 mmol, 0.2 eq.) were added in one portion under nitrogen at 25°C. The mixture was stirred at 25°C for 16 hours. The mixture was concentrated under reduced pressure at 45°C. The residue was purified by silica gel column chromatography (petroleum ether/ethyl acetate = 20/1 to 8/1) to give methyl 4-[bis(tert-butoxycarbonyl)amino]thiazole-5-carboxylate (850 mg) as a yellow oil.
工程2

エタノール(20mL)およびテトラヒドロフラン(10mL)中のメチル4-[ビス(tert-ブトキシカルボニル)アミノ]チアゾール-5-カルボキシレート(850mg、2.37mmol、1当量)の溶液に、水(1M、30mL)中の水酸化ナトリウムを25℃で添加し、混合物を70℃で3時間撹拌した。反応混合物を25℃に冷却し、pHを2M塩酸で4~5に調整し、水相を酢酸エチル(30mL×3)で抽出した。合わせた有機層をブライン(20mL×2)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、真空下で濃縮させた。化合物4-(tert-ブトキシカルボニルアミノ)チアゾール-5-カルボン酸(620mg、粗製)を白色の固体として得た。 To a solution of methyl 4-[bis(tert-butoxycarbonyl)amino]thiazole-5-carboxylate (850 mg, 2.37 mmol, 1 equiv.) in ethanol (20 mL) and tetrahydrofuran (10 mL) was added sodium hydroxide in water (1 M, 30 mL) at 25 °C, and the mixture was stirred at 70 °C for 3 h. The reaction mixture was cooled to 25 °C, the pH was adjusted to 4-5 with 2 M hydrochloric acid, and the aqueous phase was extracted with ethyl acetate (30 mL x 3). The combined organic layers were washed with brine (20 mL x 2), dried over anhydrous sodium sulfate, filtered, and concentrated under vacuum. Compound 4-(tert-butoxycarbonylamino)thiazole-5-carboxylic acid (620 mg, crude) was obtained as a white solid.
工程3

アセトニトリル(30mL)中の4-(tert-ブトキシカルボニルアミノ)チアゾール-5-カルボン酸(600mg、2.46mmol、1当量)およびリン酸カリウム(521mg、2.46mmol、1当量)の混合物に、三臭化テトラブチアンモニウム(2.37g、4.91mmol、2当量)を窒素下で、30℃において一度に添加した。混合物を50℃で16時間撹拌した。混合物を20℃に冷却し、飽和チオ硫酸ナトリウム水溶液(20mL)および飽和炭酸ナトリウム水溶液(20mL)の混合物に注ぎいれた。水相を酢酸エチル(40mL×2)で抽出した。合わせた有機相を生理食塩水(30mL)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、真空中で濃縮した。残渣を、シリカゲルカラムクロマトグラフィー(石油エーテル/酢酸エチル=30/1~8/1)によって精製して、tert-ブチルN-(5-ブロモチアゾール-4-イル)カルバメート(460mg)を黄色の固体として得た。 To a mixture of 4-(tert-butoxycarbonylamino)thiazole-5-carboxylic acid (600 mg, 2.46 mmol, 1 eq.) and potassium phosphate (521 mg, 2.46 mmol, 1 eq.) in acetonitrile (30 mL), tetrabutylammonium tribromide (2.37 g, 4.91 mmol, 2 eq.) was added in one portion under nitrogen at 30 °C. The mixture was stirred at 50 °C for 16 h. The mixture was cooled to 20 °C and poured into a mixture of saturated aqueous sodium thiosulfate (20 mL) and saturated aqueous sodium carbonate (20 mL). The aqueous phase was extracted with ethyl acetate (40 mL x 2). The combined organic phase was washed with saline (30 mL), dried over anhydrous sodium sulfate, filtered, and concentrated in vacuo. The residue was purified by silica gel column chromatography (petroleum ether/ethyl acetate = 30/1 to 8/1) to obtain tert-butyl N-(5-bromothiazol-4-yl)carbamate (460 mg) as a yellow solid.
工程4

水(3mL)およびジオキサン(18mL)中のtert-ブチルN-[(1R)-1-[4-(4,4,5,5-テトラメチル-1,3,2-ジオキサボロラン-2-イル)フェニル]エチル]カルバメート(572mg、1.65mmol、1当量)、炭酸カリウム(455mg、3.30mmol、2当量)、およびtert-ブチルN-(5-ブロモチアゾール-4-イル)カルバメート(460mg、1.65mmol、1当量)の混合物に、[1,1’-ビス(ジフェニルホスフィノ)フェロセン]ジクロロパラジウム](96mg、0.13mmol、0.08当量)を窒素下で、20℃において一度に添加した。混合物を90℃で2時間撹拌した。混合物を20℃に冷却し、氷水(w/w=1/1、20mL)に注ぎ入れ、10分間撹拌した。水相を酢酸エチル(30mL×3)で抽出した。合わせた有機相をブライン(10mL×2)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、真空中で濃縮した。残渣を、シリカゲルクロマトグラフィー(石油エーテル/酢酸エチル=30/1~5/1)によって精製して、tert-ブチルN-[5-[4-[(1S)-1-(tert-ブトキシカルボニルアミノ)エチル]フェニル]チアゾール-4-イル]カルバメート(620mg、粗製)を黄色の固体として得た。 To a mixture of tert-butyl N-[(1R)-1-[4-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)phenyl]ethyl]carbamate (572 mg, 1.65 mmol, 1 eq.), potassium carbonate (455 mg, 3.30 mmol, 2 eq.), and tert-butyl N-(5-bromothiazol-4-yl)carbamate (460 mg, 1.65 mmol, 1 eq.) in water (3 mL) and dioxane (18 mL), [1,1'-bis(diphenylphosphino)ferrocene]dichloropalladium] (96 mg, 0.13 mmol, 0.08 eq.) was added in one portion at 20°C under nitrogen. The mixture was stirred at 90°C for 2 h. The mixture was cooled to 20°C, poured into ice water (w/w=1/1, 20 mL), and stirred for 10 min. The aqueous phase was extracted with ethyl acetate (30 mL x 3). The combined organic phase was washed with brine (10 mL x 2), dried over anhydrous sodium sulfate, filtered, and concentrated in vacuum. The residue was purified by silica gel chromatography (petroleum ether/ethyl acetate = 30/1 to 5/1) to give tert-butyl N-[5-[4-[(1S)-1-(tert-butoxycarbonylamino)ethyl]phenyl]thiazol-4-yl]carbamate (620 mg, crude) as a yellow solid.
工程5

塩酸/メタノール(4M、10mL)中のtert-ブチルN-[5-[4-[(1S)-1-(tert-ブトキシカルボニルアミノ)エチル]フェニル]チアゾール-4-イル]カルバメート(620mg、1.48mmol、1当量)の溶液を、20℃で2時間撹拌した。混合物を45℃において減圧下で濃縮した。粗生成物5-[4-[(1S)-1-アミノエチル]フェニル]チアゾール-4-アミン塩酸塩(406mg、粗製)を黄色の固体として得た。 A solution of tert-butyl N-[5-[4-[(1S)-1-(tert-butoxycarbonylamino)ethyl]phenyl]thiazol-4-yl]carbamate (620 mg, 1.48 mmol, 1 equiv) in hydrochloric acid/methanol (4 M, 10 mL) was stirred at 20°C for 2 h. The mixture was concentrated under reduced pressure at 45°C. The crude product 5-[4-[(1S)-1-aminoethyl]phenyl]thiazol-4-amine hydrochloride (406 mg, crude) was obtained as a yellow solid.
上記の他の実施例について記載される手順を使用して、以下のスキームに従って、5-[4-[(1S)-1-アミノエチル]フェニル]チアゾール-4-アミンを表題化合物に変換した。

例示的な化合物638の例示的な合成
上記の他の実施例(特に、例示的な化合物238および例示的な化合物607)について記載されたものと類似の手順を使用して、以下のスキームに従って調製した。

例示的な化合物640の例示的な合成
例示的な化合物203および例示的な化合物143、ならびに上記の他の実施例について記載された手順を使用して、以下のスキームに従って調製した。

例示的な化合物642の例示的な合成
例示的な化合物143および上記の他の実施例について記載された手順を使用して、以下のスキームに従って調製した。
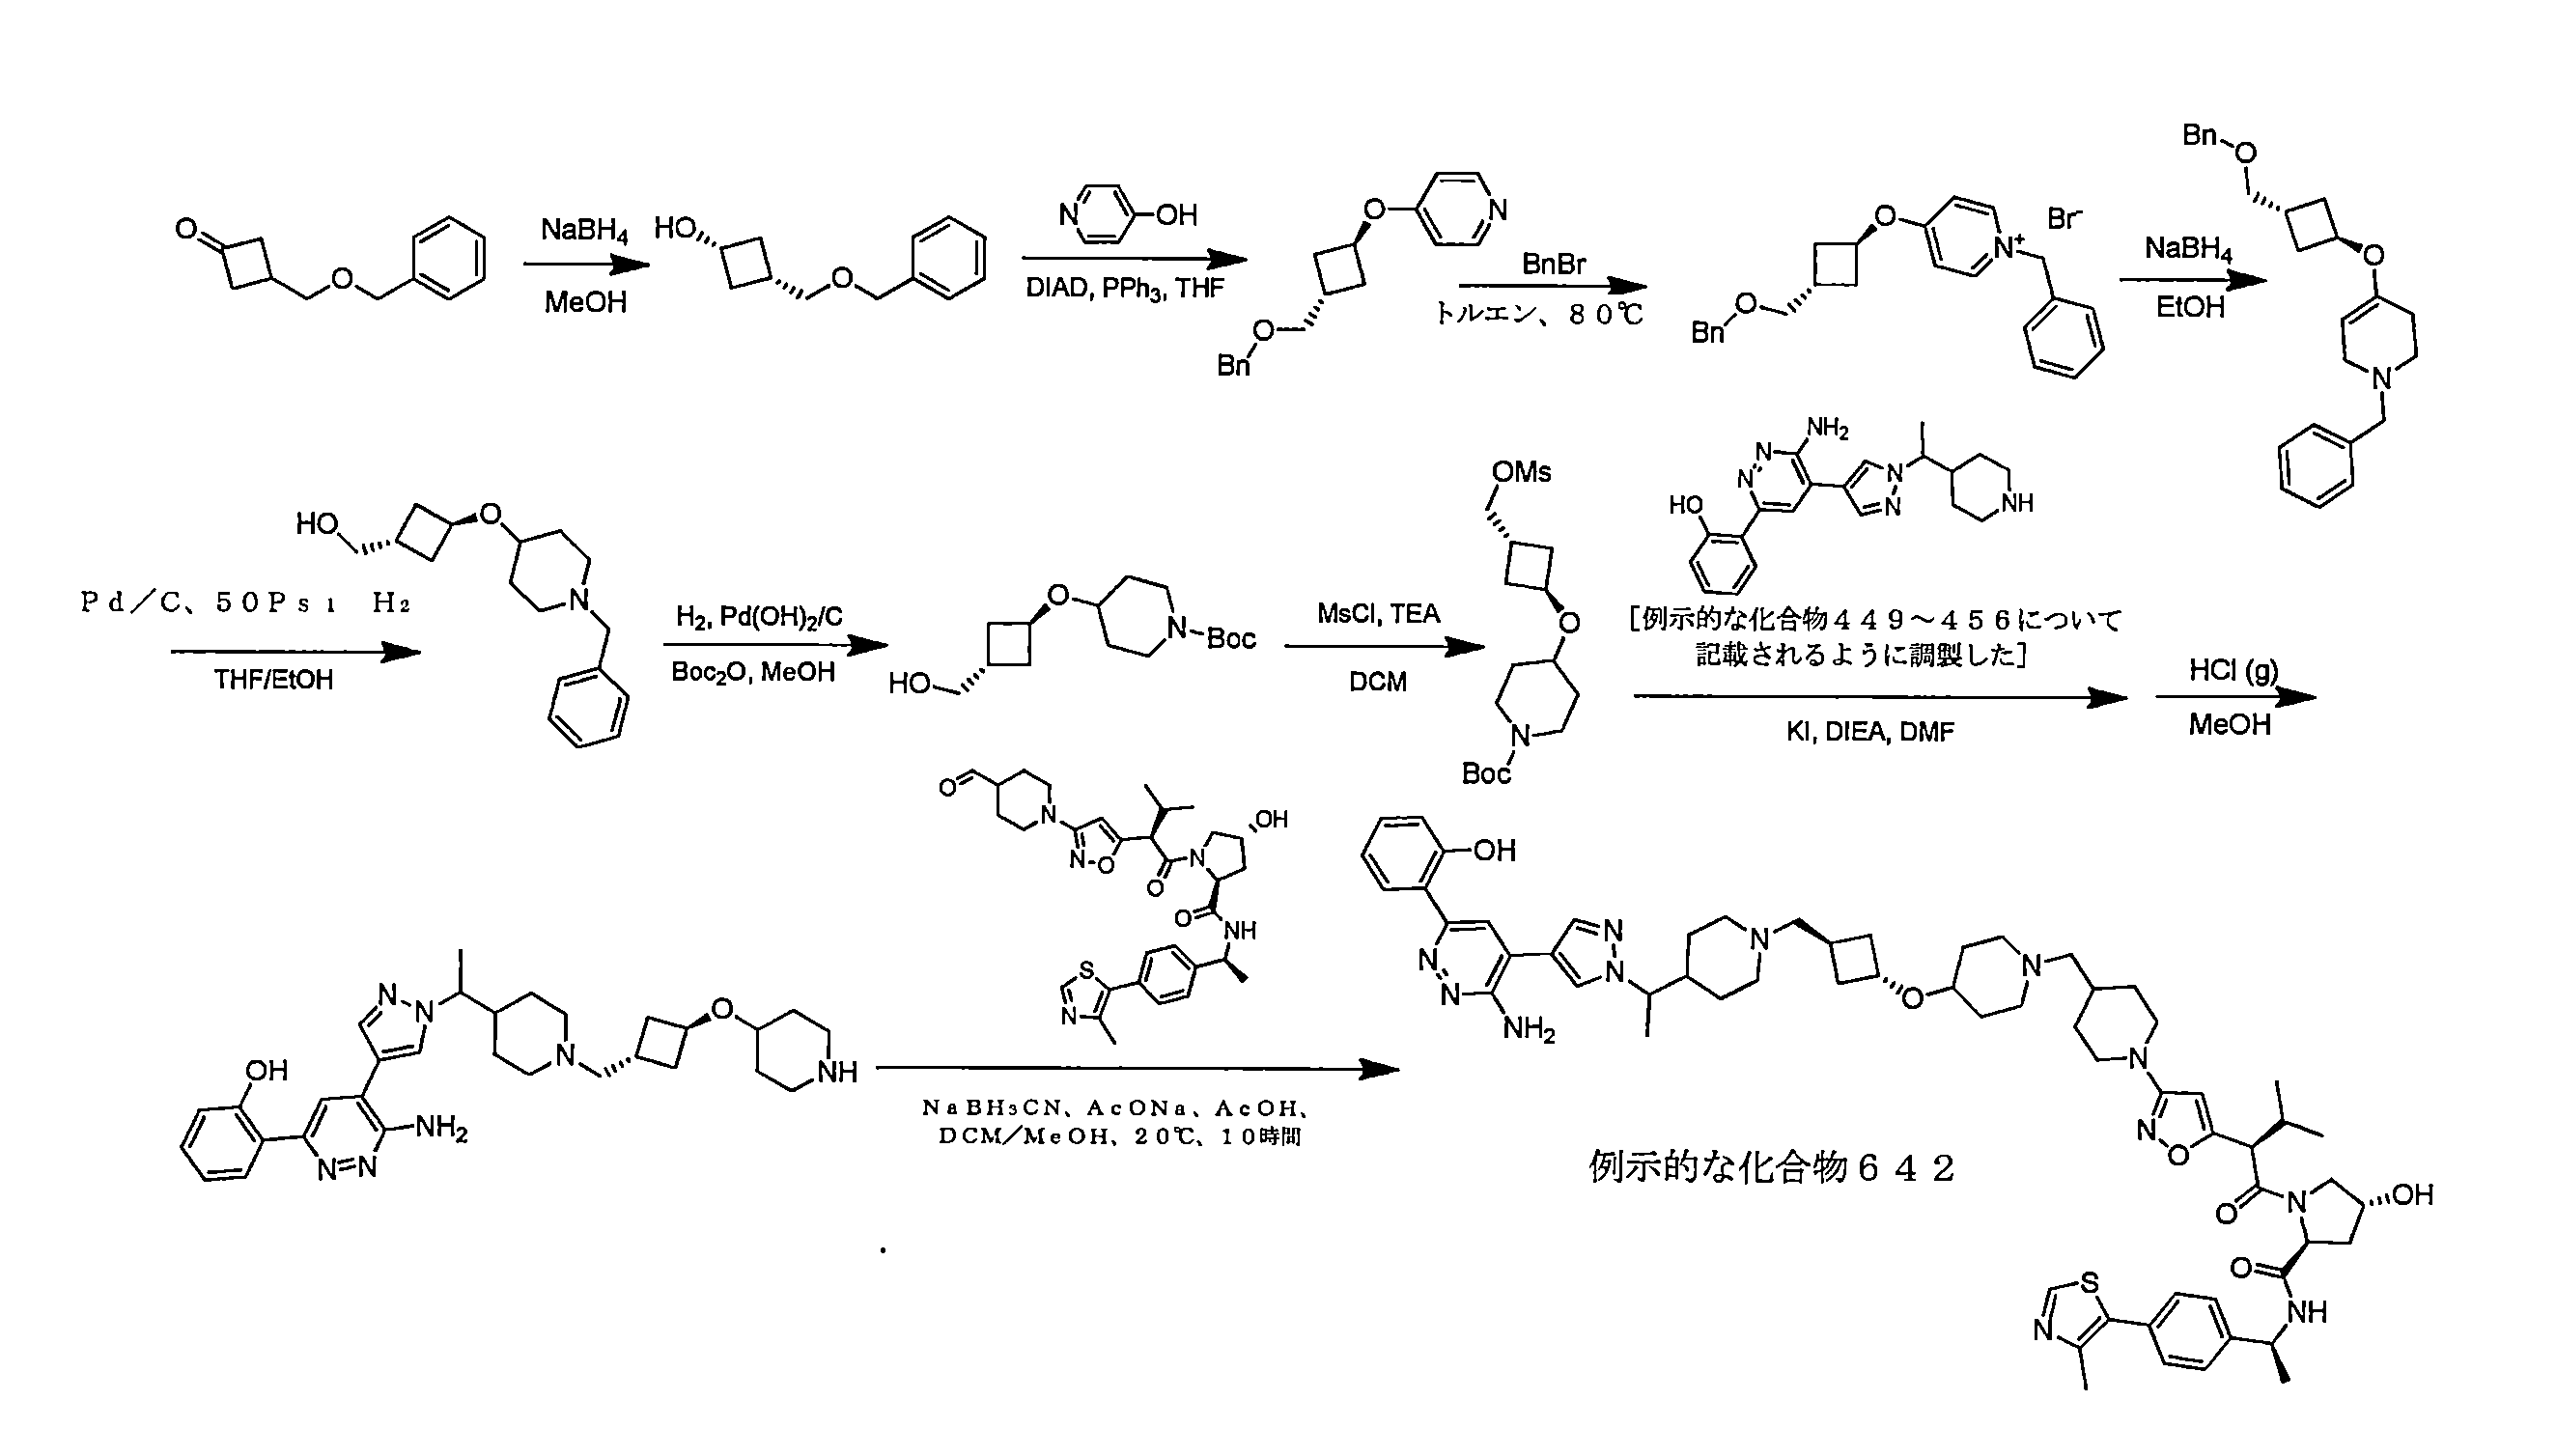
例示的な化合物664の例示的な合成
中間体2-(6-アミノ-5-(1-(1-(1-(((1s,3s)-3-(ピペリジン-4-イルオキシ)シクロブチル)メチル)ピペリジン-4-イル)エチル)-1H-ピラゾール-4-イル)ピリダジン-3-イル)フェノールを、上記の他の実施例について記載されたものと類似した手順を使用して、以下のスキームに従って調製した。

2-(6-アミノ-5-(1-(1-(1-(((1s,3s)-3-(ピペリジン-4-イルオキシ)シクロブチル)メチル)ピペリジン-4-イル)エチル)-1H-ピラゾール-4-イル)ピリダジン-3-イル)フェノールを、例示的な実施例143および上記の他の実施例について記載されるように、表題化合物に変換した。 2-(6-amino-5-(1-(1-(1-(((1s,3s)-3-(piperidin-4-yloxy)cyclobutyl)methyl)piperidin-4-yl)ethyl)-1H-pyrazol-4-yl)pyridazin-3-yl)phenol was converted to the title compound as described for illustrative Example 143 and other examples above.
例示的な化合物665の例示的な合成
中間体2-(6-アミノ-5-(1-(1-(1-((1s,3s)-3-(ピペリジン-4-イルオキシ)シクロブチル)ピペリジン-4-イル)エチル)-1H-ピラゾール-4-イル)ピリダジン-3-イル)フェノールを、以下のスキームに従って調製した。

2-(6-アミノ-5-(1-(1-(1-((1s,3s)-3-(ピペリジン-4-イルオキシ)シクロブチル)ピペリジン-4-イル)エチル)-1H-ピラゾール-4-イル)ピリダジン-3-イル)フェノールを、例示的な実施例143および上記の他の実施例について記載されるように、表題化合物に変換した。 2-(6-amino-5-(1-(1-(1-((1s,3s)-3-(piperidin-4-yloxy)cyclobutyl)piperidin-4-yl)ethyl)-1H-pyrazol-4-yl)pyridazin-3-yl)phenol was converted to the title compound as described for illustrative Example 143 and other examples above.
例示的な化合物678の例示的な合成
中間体(2S,4R)-N-((S)-1-(4-エチニルフェニル)エチル)-1-((R)-2-(3-(4-ホルミルピペリジン-1-イル)イソキサゾール-5-イル)-3-メチルブタノイル)-4-ヒドロキシピロリジン-2-カルボキサミドを、以下のスキームに従って調製した。

(2S,4R)-N-((S)-1-(4-エチニルフェニル)エチル)-1-((R)-2-(3-(4-ホルミルピペリジン-1-イル)イソキサゾール-5-イル)-3-メチルブタノイル)-4-ヒドロキシピロリジン-2-カルボキサミドを、上記の例示的な化合物143について記載された手順を使用して、表題化合物に変換した。 (2S,4R)-N-((S)-1-(4-ethynylphenyl)ethyl)-1-((R)-2-(3-(4-formylpiperidin-1-yl)isoxazol-5-yl)-3-methylbutanoyl)-4-hydroxypyrrolidine-2-carboxamide was converted to the title compound using the procedure described for exemplary compound 143 above.
例示的な化合物692の例示的な合成
工程1

アセトニトリル(2000mL)中の3-ベンジルオキシシクロブタノール(100g、561.08mmol、1当量)およびベンジル4-オキソピペリジン-1-カルボキシレート(143.97g、617.19mmol、123.1mL、1.1当量)の混合物を脱気し、窒素で3回パージした。次いで、混合物に、クロロ(ジメチル)シラン(53.09g、561.08mmol、1当量)を0℃で添加した。混合物を、窒素雰囲気下で、25℃において12時間撹拌した。反応混合物を水(2L)で希釈した。有機層を酢酸エチル(1L×2)で抽出した。合わせた有機層をブライン(500mL)で洗浄した。有機層を無水硫酸ナトリウムで乾燥させ、濾過し、真空下で濃縮して、残渣を得た。残渣を、シリカゲルクロマトグラフィー(石油エーテル/酢酸エチル=30/1~20:1)によって精製して、生成物を得た。ベンジル4-(3-ベンジルオキシシクロブトキシ)ピぺリジン-1-カルボキシレート(89g、225.04mmol)を無色の油として得た。 A mixture of 3-benzyloxycyclobutanol (100 g, 561.08 mmol, 1 eq.) and benzyl 4-oxopiperidine-1-carboxylate (143.97 g, 617.19 mmol, 123.1 mL, 1.1 eq.) in acetonitrile (2000 mL) was degassed and purged with nitrogen three times. To the mixture was then added chloro(dimethyl)silane (53.09 g, 561.08 mmol, 1 eq.) at 0° C. The mixture was stirred at 25° C. under nitrogen atmosphere for 12 hours. The reaction mixture was diluted with water (2 L). The organic layer was extracted with ethyl acetate (1 L×2). The combined organic layers were washed with brine (500 mL). The organic layer was dried over anhydrous sodium sulfate, filtered, and concentrated under vacuum to give a residue. The residue was purified by silica gel chromatography (petroleum ether/ethyl acetate=30/1 to 20:1) to give the product. Benzyl 4-(3-benzyloxycyclobutoxy)piperidine-1-carboxylate (89 g, 225.04 mmol) was obtained as a colorless oil.
工程2

エタノール(300mL)およびテトラヒドロフラン(300mL)中のベンジル4-(3-ベンジルオキシシクロブトキシ)ピペリジン-1-カルボキシレート(65g、164.35mmol、1当量)の溶液に、活性化炭素触媒上のパラジウム(6g、1.44mmol、10%純度)、活性化炭素触媒上の水酸化パラジウム(6g、8.54mmol、20%純度)、およびジ-tert-ブチルジカーボネート(53.80g、246.53mmol、56.6mL、1.5当量)を窒素雰囲気下で添加した。懸濁液を脱気し、水素で3回パージした。混合物を、水素下(50Psi)で、40℃において16時間撹拌した。反応混合物を減圧下で濾過および濃縮し、残渣を得た。粗生成物をシリカゲルクロマトグラフィー(石油エーテル/酢酸エチル=50/1、0/1)によって精製した。粗生成物tert-ブチル4-(3-ヒドロキシシクロブトキシ)ピペリジン-1-カルボキシレート(30.8g、113.5mmol)を白色の固体として得た。 To a solution of benzyl 4-(3-benzyloxycyclobutoxy)piperidine-1-carboxylate (65 g, 164.35 mmol, 1 equiv.) in ethanol (300 mL) and tetrahydrofuran (300 mL), palladium on activated carbon catalyst (6 g, 1.44 mmol, 10% purity), palladium hydroxide on activated carbon catalyst (6 g, 8.54 mmol, 20% purity), and di-tert-butyl dicarbonate (53.80 g, 246.53 mmol, 56.6 mL, 1.5 equiv.) were added under nitrogen atmosphere. The suspension was degassed and purged with hydrogen three times. The mixture was stirred under hydrogen (50 Psi) at 40 °C for 16 h. The reaction mixture was filtered and concentrated under reduced pressure to give a residue. The crude product was purified by silica gel chromatography (petroleum ether/ethyl acetate = 50/1, 0/1). The crude product, tert-butyl 4-(3-hydroxycyclobutoxy)piperidine-1-carboxylate (30.8 g, 113.5 mmol), was obtained as a white solid.
工程3

トルエン(40mL)中のtert-ブチル4-(3-ヒドロキシシクロブトキシ)ピペリジン-1-カルボキシレート(5g、18.43mmol、1当量)、4-ブロモピリジン-2-オール(3.85g、22.11mmol、1.2当量)、トリフェニルホスフィン(7.25g、27.64mmol、1.5当量)、およびジイソプロピルアゾジカルボキシレート(5.59g、27.64mmol、5.4mL、1.5当量)を脱気し、窒素で3回パージし、次いで、混合物を窒素雰囲気下で、110℃において12時間撹拌した。反応混合物を20℃に冷却し、次いで、濾過し、減圧下で濃縮して、残渣を得た。残渣を、シリカゲルクロマトグラフィー(石油エーテル/酢酸エチル=50/1~10/1)によって精製して、生成物を得た。tert-ブチル4-[3-[(4-ブロモ-2-ピリジル)オキシ]シクロブトキシ]ピペリジン-1-カルボキシレート(7g、16.38mmol)を淡い黄色の油として得た。 tert-Butyl 4-(3-hydroxycyclobutoxy)piperidine-1-carboxylate (5 g, 18.43 mmol, 1 eq.), 4-bromopyridin-2-ol (3.85 g, 22.11 mmol, 1.2 eq.), triphenylphosphine (7.25 g, 27.64 mmol, 1.5 eq.), and diisopropyl azodicarboxylate (5.59 g, 27.64 mmol, 5.4 mL, 1.5 eq.) in toluene (40 mL) was degassed and purged with nitrogen three times, and then the mixture was stirred under nitrogen atmosphere at 110° C. for 12 hours. The reaction mixture was cooled to 20° C., then filtered and concentrated under reduced pressure to give a residue. The residue was purified by silica gel chromatography (petroleum ether/ethyl acetate=50/1 to 10/1) to give the product. tert-Butyl 4-[3-[(4-bromo-2-pyridyl)oxy]cyclobutoxy]piperidine-1-carboxylate (7 g, 16.38 mmol) was obtained as a pale yellow oil.
工程4
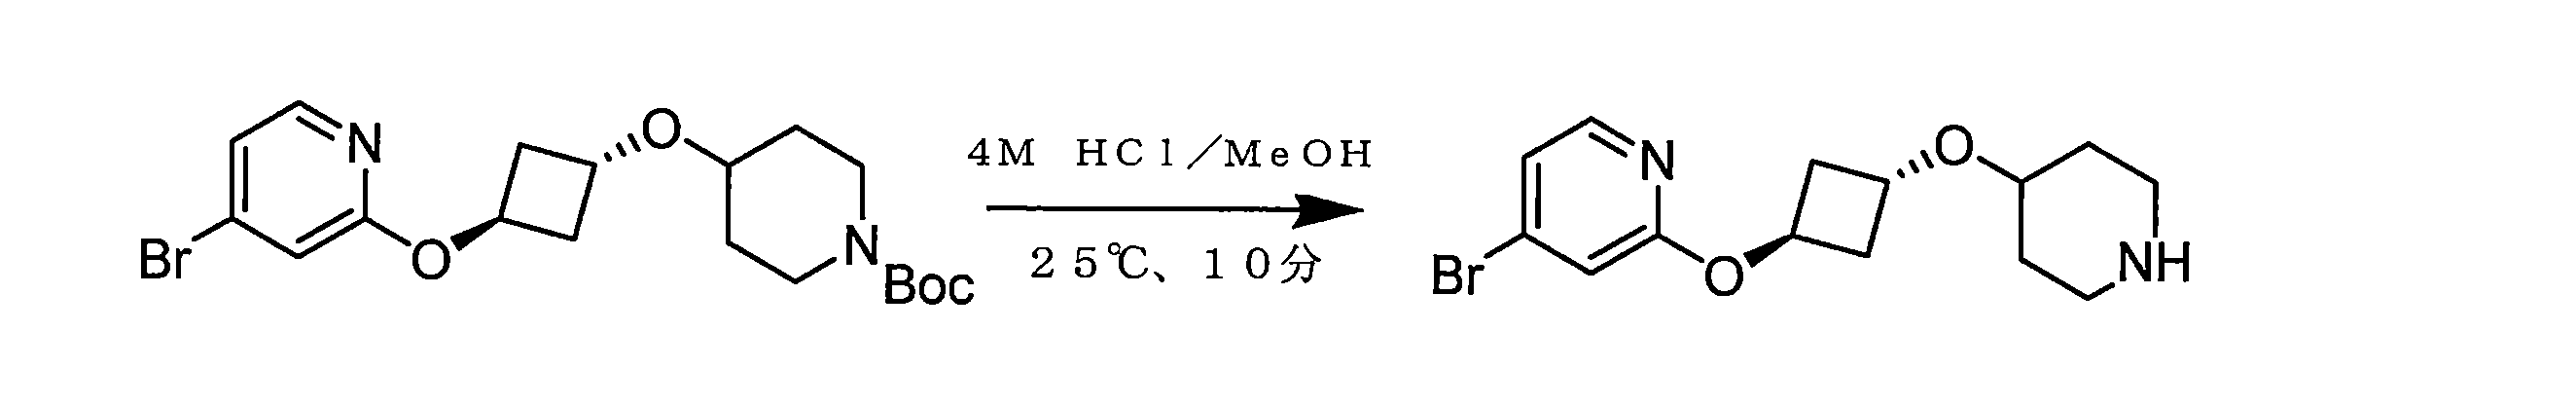
塩酸塩/メタノール(4M、25mL、10.68当量)中のtert-ブチル4-[3-[(4-ブロモ-2-ピリジル)オキシ]シクロブトキシ]ピペリジン-1-カルボキシレート(4g、9.36mmol、1当量)の溶液を、25℃で10分間撹拌した。反応溶液を真空下で濃縮して、粗生成物を得た。粗生成物をさらに精製することなく次の工程で直接使用した。4-ブロモ-2[3-(4-ピペリジルオキシ)シクロブトキシ]ピリジン塩酸塩(3.2g、8.80mmol)を淡い黄色の油として得た。 A solution of tert-butyl 4-[3-[(4-bromo-2-pyridyl)oxy]cyclobutoxy]piperidine-1-carboxylate (4 g, 9.36 mmol, 1 eq.) in hydrochloride/methanol (4 M, 25 mL, 10.68 eq.) was stirred at 25°C for 10 min. The reaction solution was concentrated under vacuum to give the crude product, which was used directly in the next step without further purification. 4-Bromo-2[3-(4-piperidyloxy)cyclobutoxy]pyridine hydrochloride (3.2 g, 8.80 mmol) was obtained as a pale yellow oil.
工程5

N,N-ジメチルホルムアミド(40mL)中の4-ブロモ-2-[3-(4-ピペリジルオキシ)シクロブトキシ]ピリジン(3.2g、8.80mmol、1当量、塩酸塩)の溶液に、トリエチルアミン(5.34g、52.79mmol、7.4mL、6当量)および1-tert-ブトキシカルボニルピペリジン-4-カルボン酸(2.42g、10.56mmol、1.2当量)を添加した。添加後、混合物を20℃で2分間撹拌した。次いで、混合物に、O-(7-アザベンゾトリアゾール-1-イル)-N,N,N’,N’-テトラメチルウロニウムヘキサフルオロホスフェート(4.01g、10.56mmol、1.2当量)を0℃で添加し、混合物を窒素下で、20℃において1時間撹拌した。反応混合物を水(40mL)で希釈し、酢酸エチル(100mL×2)で抽出した。合わせた有機層を飽和ブライン(100mL)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、真空中で濃縮した。残渣を、分取HPLC(カラム:Phenomenex luna c18 250mm*100mm*10um;移動相:[水(0.225%FA)-ACN];B%:40%~70%、20分)によって精製した。化合物tert-ブチル4-[4-[3-[(4-ブロモ-2-ピリジル)オキシ]シクロブトキシ]ピペリジン-1-カルボニル]ピペリジン-1-カルボキシレート(4.5g、8.36mmol)を白色の固体として得た。 To a solution of 4-bromo-2-[3-(4-piperidyloxy)cyclobutoxy]pyridine (3.2 g, 8.80 mmol, 1 eq., hydrochloride salt) in N,N-dimethylformamide (40 mL) was added triethylamine (5.34 g, 52.79 mmol, 7.4 mL, 6 eq.) and 1-tert-butoxycarbonylpiperidine-4-carboxylic acid (2.42 g, 10.56 mmol, 1.2 eq.). After addition, the mixture was stirred at 20°C for 2 minutes. To the mixture was then added O-(7-azabenzotriazol-1-yl)-N,N,N',N'-tetramethyluronium hexafluorophosphate (4.01 g, 10.56 mmol, 1.2 eq.) at 0°C, and the mixture was stirred under nitrogen at 20°C for 1 hour. The reaction mixture was diluted with water (40 mL) and extracted with ethyl acetate (100 mL x 2). The combined organic layers were washed with saturated brine (100 mL), dried over anhydrous sodium sulfate, filtered, and concentrated in vacuum. The residue was purified by preparative HPLC (column: Phenomenex luna c18 250 mm * 100 mm * 10 um; mobile phase: [water (0.225% FA)-ACN]; B%: 40%-70%, 20 min). The compound tert-butyl 4-[4-[3-[(4-bromo-2-pyridyl)oxy]cyclobutoxy]piperidine-1-carbonyl]piperidine-1-carboxylate (4.5 g, 8.36 mmol) was obtained as a white solid.
工程6
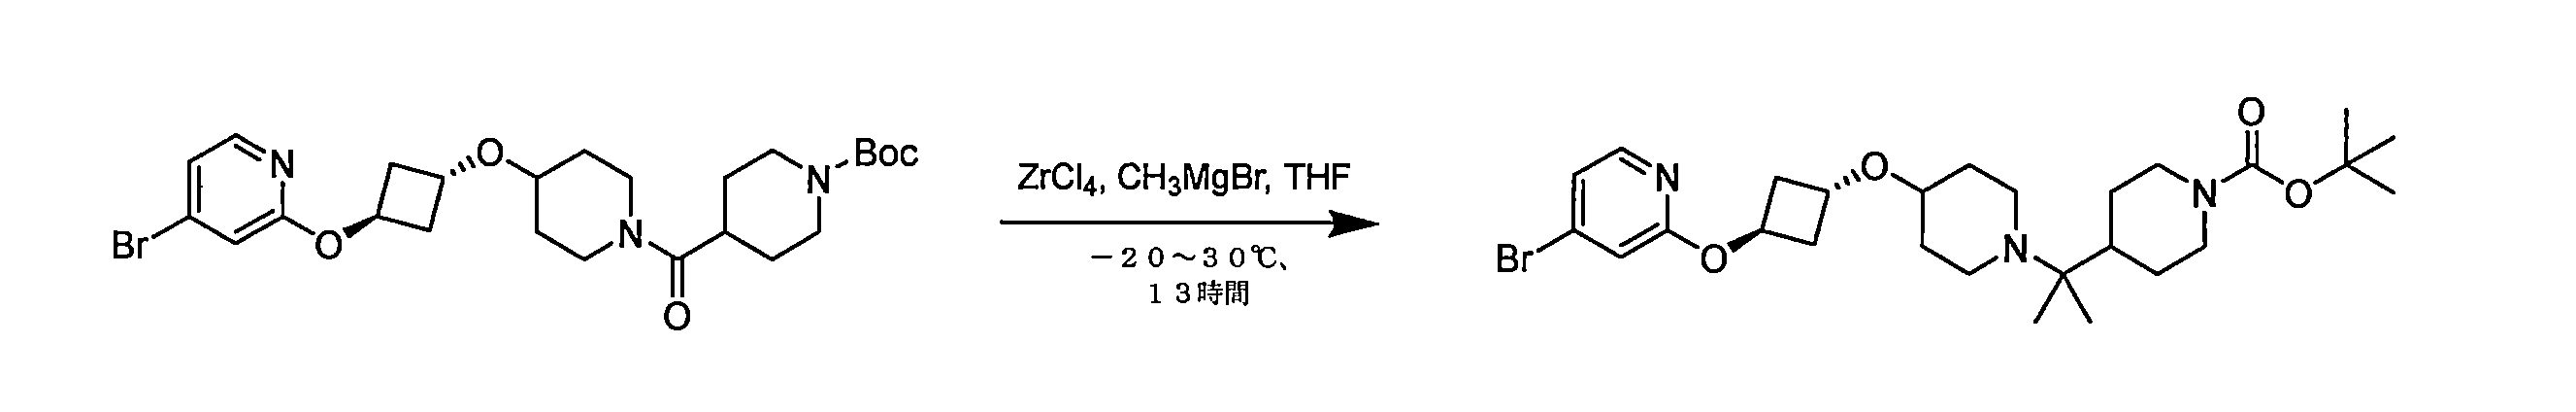
テトラヒドロフラン(70mL)中のtert-ブチル4-[4-[3-[(4-ブロモ-2-ピリジル)オキシ]シクロブトキシ]ピペリジン-1-カルボニル]ピペリジン-1-カルボキシレート(3.5g、6.50mmol、1当量)の溶液に、塩化ジルコニウム(IV)(3.03g、13.0mmol、1.1mL、2当量)を-20℃で添加し、混合物を-20℃で1時間撹拌した。次いで、臭化メチルマグネシウム(3M、13mL、6当量)を-10℃でゆっくり混合物に添加した。混合物を30℃で12時間撹拌した。反応混合物を、飽和塩化アンモニウム溶液(100mL)によって希釈し、酢酸エチル(200mL×2)で抽出した。合わせた有機層を飽和ブライン(100mL)で洗浄し、無水硫酸ナトリウムで乾燥させ、濾過し、真空中で濃縮した。残渣を、分取HPLC(カラム:Phenomenex Synergi Max-RP 250*50mm*10um;移動相:[水(0.1%TFA)-ACN];B%:25%~50%、20分)によって精製した。化合物tert-ブチル4-[1-[4-[3-[(4-ブロモ-2-ピリジル)オキシ]シクロブトキシ]-1-ピペリジル]-1-メチル-エチル]ピペリジン-1-カルボキシレート(1g、1.81mmol、収率27%)を淡い黄色の油として得た。 To a solution of tert-butyl 4-[4-[3-[(4-bromo-2-pyridyl)oxy]cyclobutoxy]piperidine-1-carbonyl]piperidine-1-carboxylate (3.5 g, 6.50 mmol, 1 eq.) in tetrahydrofuran (70 mL), zirconium chloride (IV) (3.03 g, 13.0 mmol, 1.1 mL, 2 eq.) was added at -20 ° C. and the mixture was stirred at -20 ° C. for 1 h. Then, methylmagnesium bromide (3 M, 13 mL, 6 eq.) was added slowly to the mixture at -10 ° C. The mixture was stirred at 30 ° C. for 12 h. The reaction mixture was diluted by saturated ammonium chloride solution (100 mL) and extracted with ethyl acetate (200 mL x 2). The combined organic layers were washed with saturated brine (100 mL), dried over anhydrous sodium sulfate, filtered, and concentrated in vacuum. The residue was purified by preparative HPLC (column: Phenomenex Synergi Max-RP 250*50mm*10um; mobile phase: [water (0.1% TFA)-ACN]; B%: 25%-50%, 20min). The compound tert-butyl 4-[1-[4-[3-[(4-bromo-2-pyridyl)oxy]cyclobutoxy]-1-piperidyl]-1-methyl-ethyl]piperidine-1-carboxylate (1g, 1.81mmol, 27% yield) was obtained as a pale yellow oil.
例示的な化合物143、例示的な化合物616、および他の実施例について記載されるものと類似した手順を使用して、以下のスキームに従って、tert-ブチル4-[1-[4-[3-[(4-ブロモ-2-ピリジル)オキシ]シクロブトキシ]-1-ピペリジル]-1-メチル-エチル]ピペリジン-1-カルボキシレートを表題化合物に変換した。

例示的な化合物698の例示的な合成
例示的な化合物638および例示的な化合物640に記載されるものと類似した手順を使用して、以下のスキームに従って調製した。
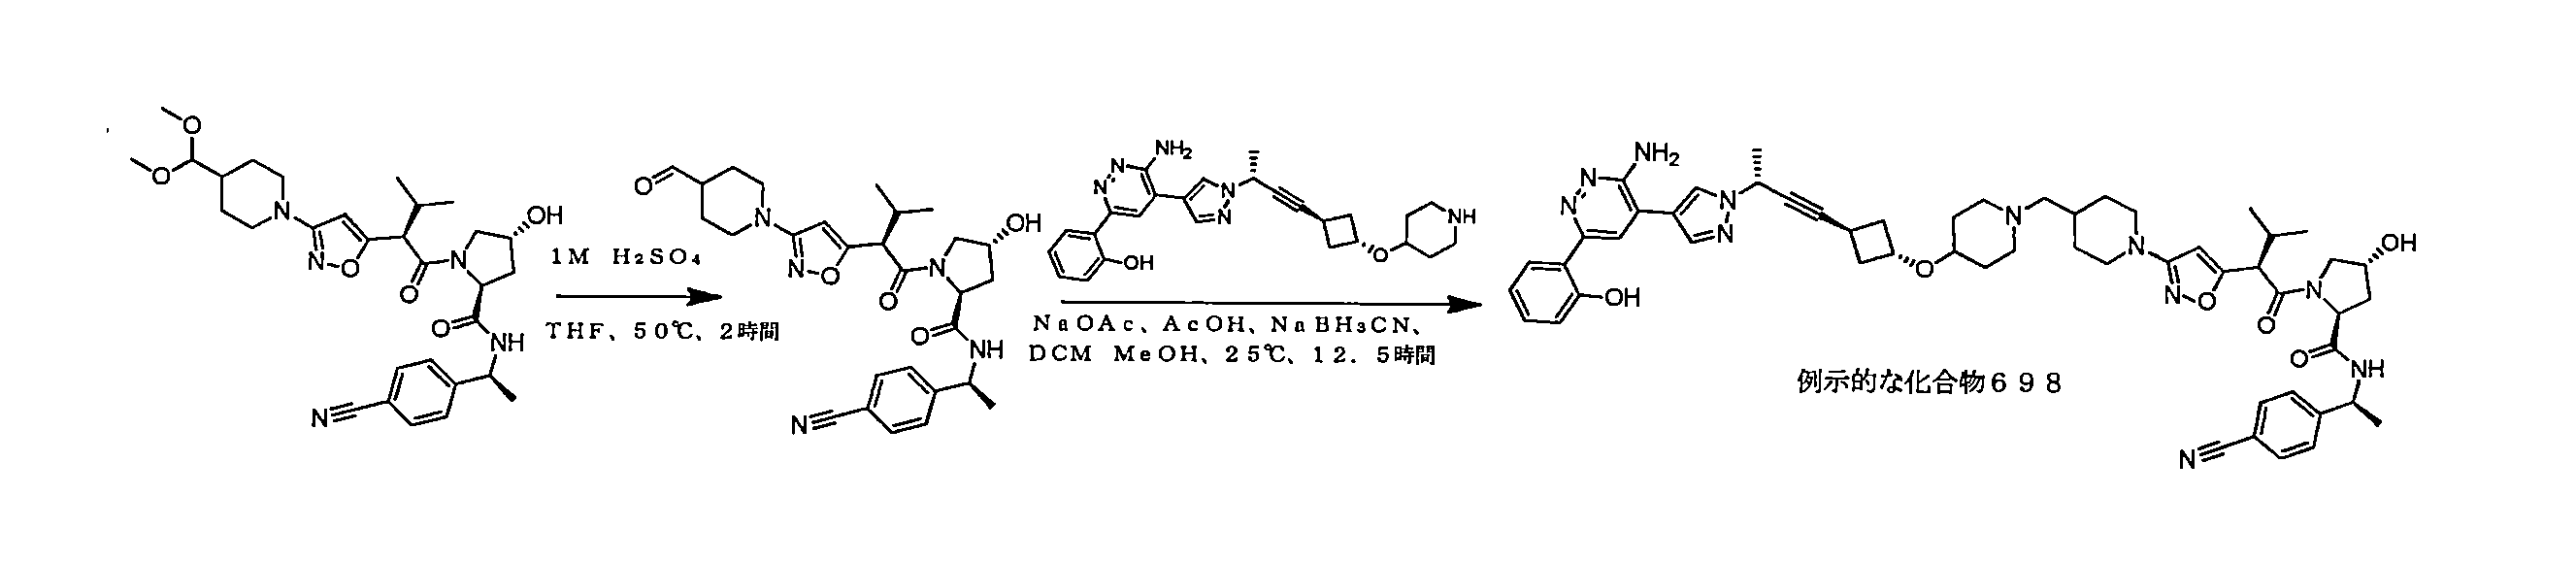
例示的な化合物707の例示的な合成
上記の他の実施例について記載された手順を使用して、以下のスキームに従って調製した。
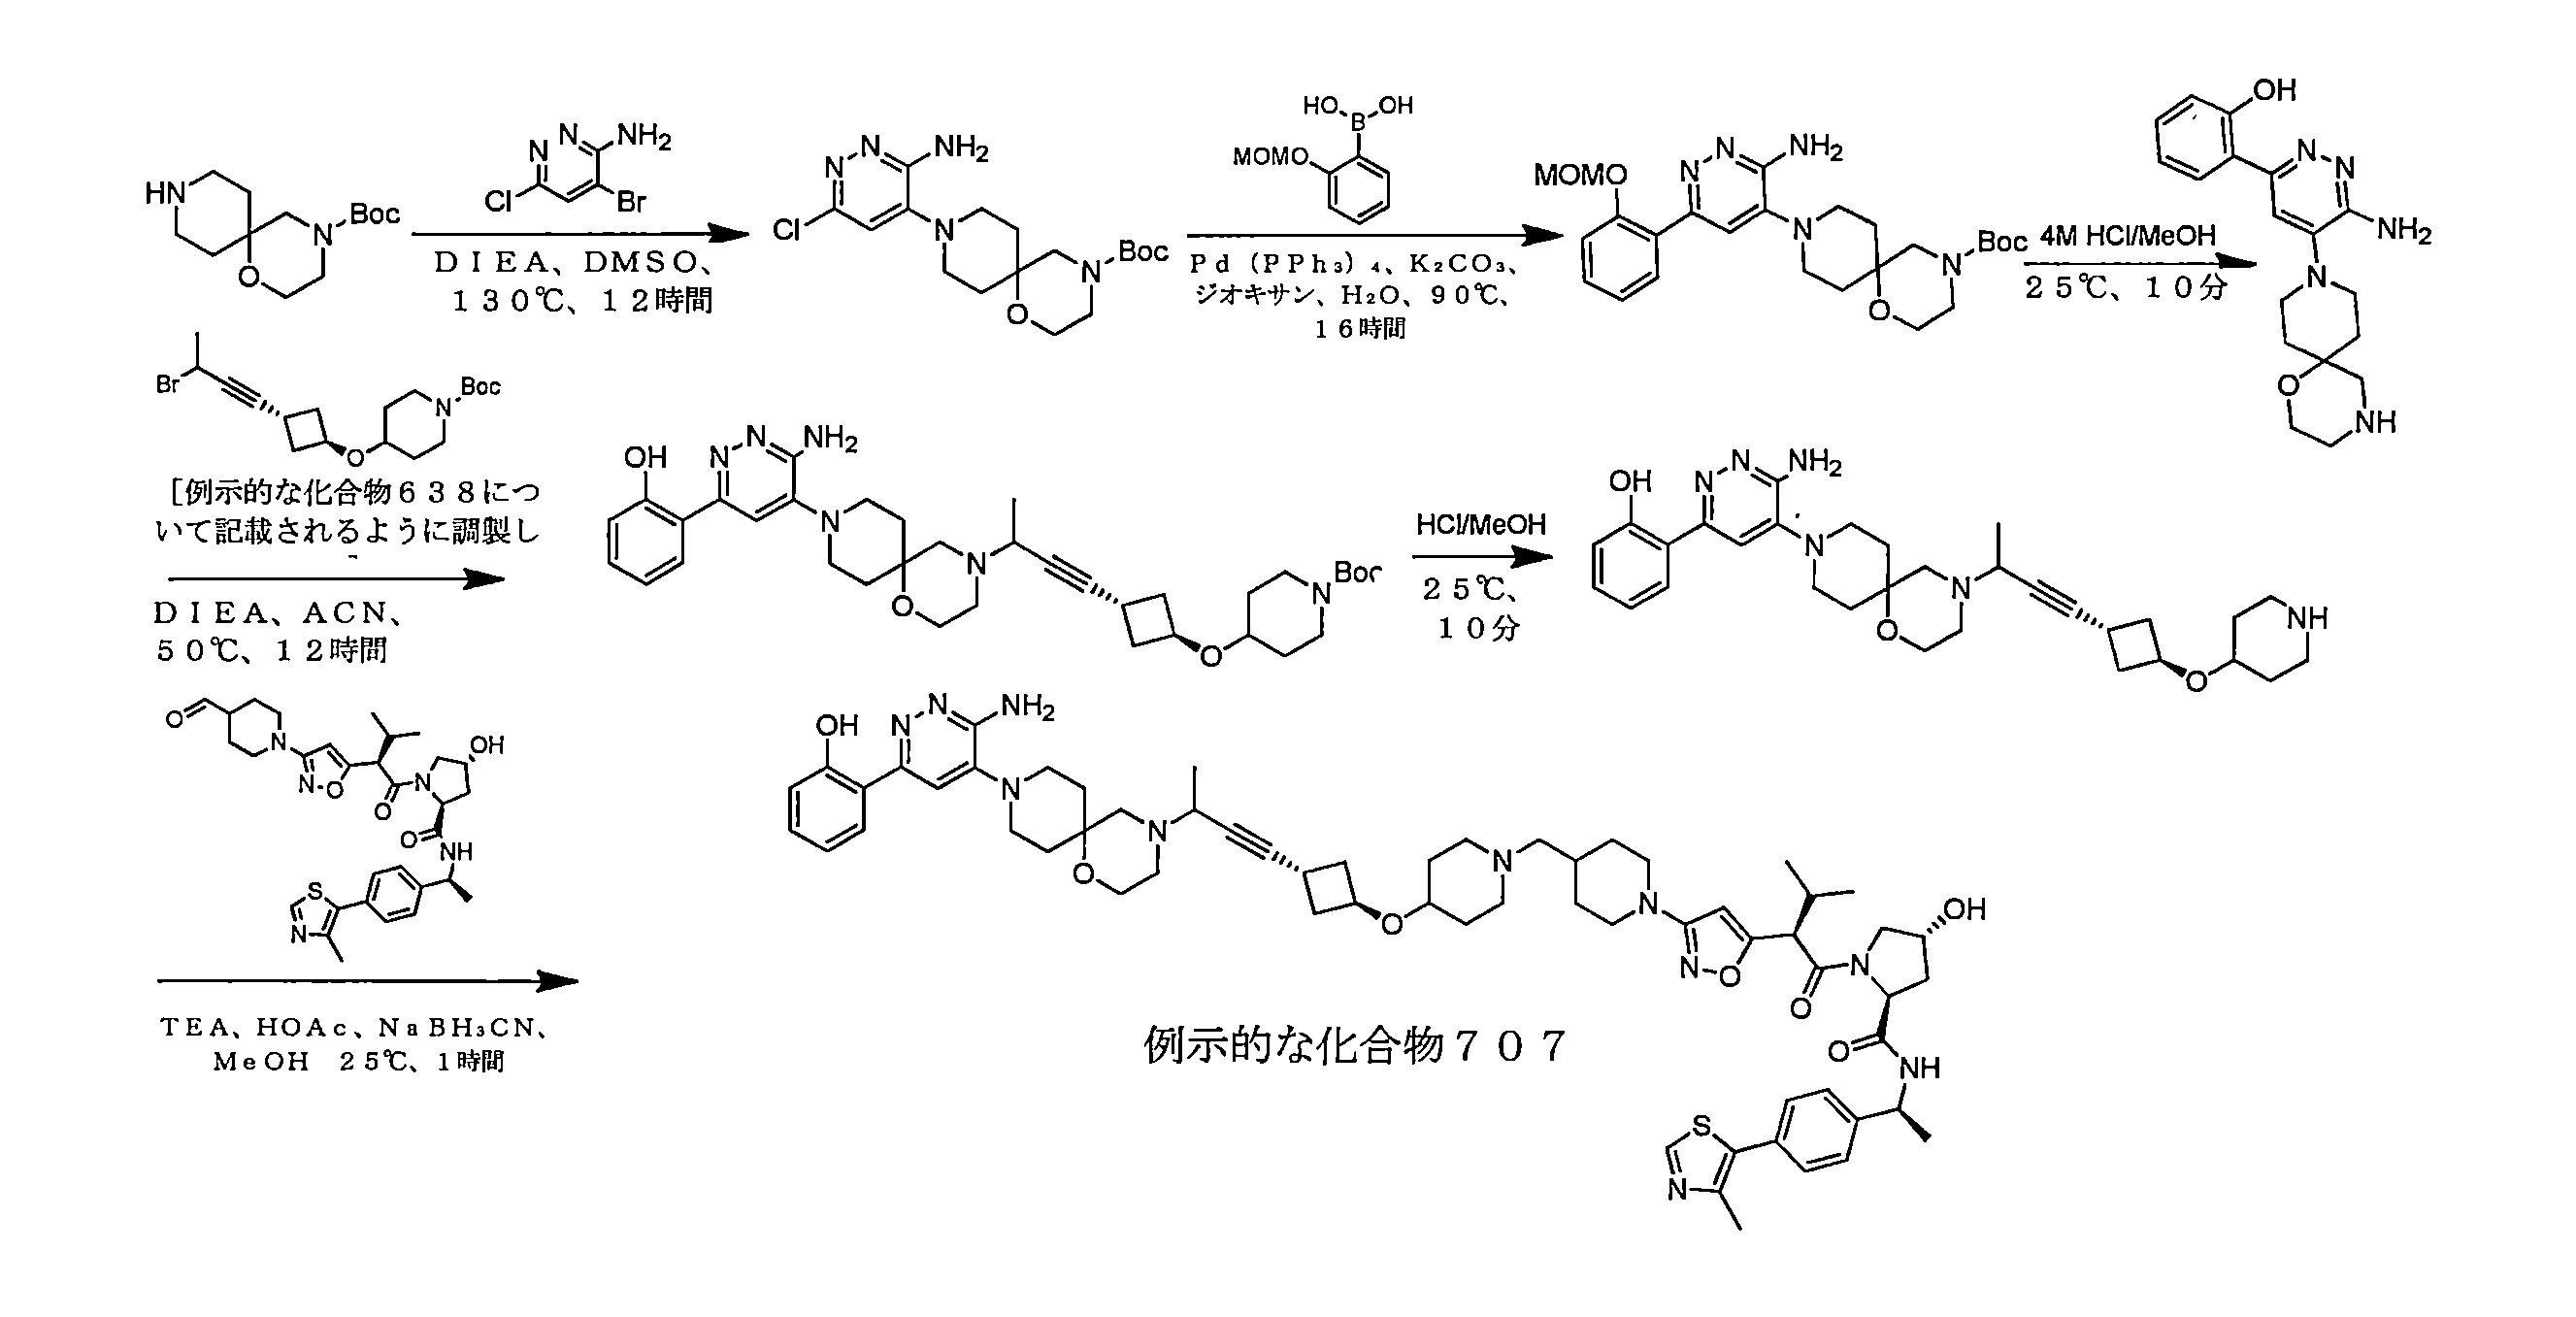
タンパク質レベルの制御
本明細書はさらに、細胞を用いたタンパク質レベルの制御方法を提供する。この方法は、特定の標的タンパク質と相互作用することが知られている、本明細書に記載される化合物の使用に基づいており、それにより標的タンパク質のインビボ分解が、好ましくは特定の治療利益に対して、生物系中のタンパク質量の制御をもたらす。
The present specification further provides a method for controlling protein levels using cells. The method is based on the use of compounds described herein that are known to interact with specific target proteins, whereby the in vivo degradation of the target protein leads to the control of the amount of protein in a biological system, preferably for a specific therapeutic benefit.
以下の実施例は、本開示の解説を補完するために使用されるが、本開示を限定するものとしては決してみなされない。 The following examples are used to complement the explanation of this disclosure, but are in no way to be construed as limiting this disclosure.
本開示の具体的な実施形態
本開示は、以下の具体的な実施形態を包含する。以下のこれら実施形態は、以下に続く実施形態において列挙される特性のすべてを規定されるように含み得る。適切である場合、以下の実施形態はさらに、以下に続く実施形態において列挙される特性を包括的に、または選択的に含み得る(例え第八の実施形態は、第一の実施形態において列挙される特性を列挙されるように含み得、および/または第二から第七の実施形態のいずれかの特性を含み得る。
Specific Embodiments of the Disclosure The disclosure includes the following specific embodiments, which may include all of the features recited in the embodiments that follow, as specified. Where appropriate, the embodiments may further include, inclusively or in the alternative, the features recited in the embodiments that follow (e.g., the eighth embodiment may include the features recited in the first embodiment, as recited, and/or may include any of the features of the second through seventh embodiments).
ある特定の実施形態では、説明は、その塩、プロドラッグ、多形体、類似体、誘導体、および重水素化形態を含む、以下の例示的なSMARCA2(すなわち、BRAHMAまたはBRM)PROTAC分子(表1A、表1B、表1C、表1D、表2A、表2B、表2C、および表2Dの化合物1~765)を提供する。ある特定の実施形態では、説明は、以下の化学構造:PTM-L-ULMを有する二官能性化合物であって、ULMが、フォンヒッペル・リンダウE3ユビキチンリガーゼに結合する小分子E3ユビキチンリガーゼ結合部分であり、Lが、ULMとPTMとを連結する結合または化学結合部分であり、PTMが、式I、II、III、IVa、またはIVbで表される化学構造を有するSMARCA2タンパク質標的化部分を含む小分子であり、

WPTM1は、任意に置換された5~6員アリールまたはヘテロアリール環(例えば、ヒドロキシ、ハロゲン、アルコキシ、アルキル、ハロアルキル、アミノ、アルキルアミノ、およびシアノから選択される0、1、2、または3個の置換基で置換された5~6員アリールまたはヘテロアリール)であり、
WPTM2が、任意に置換された5~6員アリールまたはヘテロアリール環(例えば、ヒドロキシ、ハロゲン、アルコキシ、アルキル、ハロアルキル、アミノ、アルキルアミノ、およびシアノから選択される0、1、2、または3個の置換基で置換された5~6員アリールまたはヘテロアリール)であり、
WPTM3は、存在しないか、または任意に置換された5~6員アリールもしくはヘテロアリール環(例えば、ヒドロキシ、ハロゲン、アルコキシ、アルキル、ハロアルキル、アミノ、アルキルアミノ、およびシアノから選択される0、1、2、もしくは3個の置換基で置換された5~6員アリールもしくはヘテロアリール)、または任意に置換された4~9シクロアルキルもしくはヘテロシクリル、任意に置換された架橋ビシクロアルキルおよび架橋ビヘテロシクリル(例えば、ヒドロキシ、ハロゲン、アルコキシ、アルキル、ハロアルキル、アミノ、アルキルアミノ、およびシアノから選択される0、1、もしくは2個の置換基で置換された4~9シクロアルキルもしくはヘテロシクリル)であり、
WPTM4は、WPTM2環で縮合された任意に置換された5~7シクロアルキルまたはヘテロシクリル(例えば、ヒドロキシ、ハロゲン、アルコキシ、アルキル、ハロアルキル、アミノ、アルキルアミノ、およびシアノから選択される0、1、2、または3個の置換基で置換された5~7シクロアルキルまたはヘテロシクリル)であり、
WPTM5は、存在しないか、または任意に置換された5~6員アリールもしくはヘテロアリール環(例えば、ヒドロキシ、ハロゲン、アルコキシ、アルキル、ハロアルキル、アミノ、アルキルアミノ、およびシアノから選択される0、1、または2個の置換基で置換された5~6員アリールまたはヘテロアリール)であり、
WPTM6およびWPTM7は、独立して、任意に4~7シクロアルキルまたはヘテロシクリル(例えば、各々が独立して、ヒドロキシ、ハロゲン、アルコキシ、アルキル、ハロアルキル、アミノ、アルキルアミノ、およびシアノから選択される0、1、または2個の置換基で置換された4~7シクロアルキルまたはヘテロシクリルである)であり、WPTM6およびWPTM7の環は、スピロ連結を介して縮合または結合され、

WPTM1 is an optionally substituted 5-6 membered aryl or heteroaryl ring (e.g., a 5-6 membered aryl or heteroaryl substituted with 0, 1, 2, or 3 substituents selected from hydroxy, halogen, alkoxy, alkyl, haloalkyl, amino, alkylamino, and cyano);
WPTM2 is an optionally substituted 5-6 membered aryl or heteroaryl ring (e.g., a 5-6 membered aryl or heteroaryl substituted with 0, 1, 2, or 3 substituents selected from hydroxy, halogen, alkoxy, alkyl, haloalkyl, amino, alkylamino, and cyano);
WPTM3 is absent or an optionally substituted 5-6 membered aryl or heteroaryl ring (e.g., a 5-6 membered aryl or heteroaryl substituted with 0, 1, 2, or 3 substituents selected from hydroxy, halogen, alkoxy, alkyl, haloalkyl, amino, alkylamino, and cyano), or an optionally substituted 4-9 cycloalkyl or heterocyclyl, optionally substituted bridged bicycloalkyl, and bridged biheterocyclyl (e.g., a 4-9 cycloalkyl or heterocyclyl substituted with 0, 1, or 2 substituents selected from hydroxy, halogen, alkoxy, alkyl, haloalkyl, amino, alkylamino, and cyano);
WPTM4 is an optionally substituted 5-7 cycloalkyl or heterocyclyl (e.g., 5-7 cycloalkyl or heterocyclyl substituted with 0, 1, 2, or 3 substituents selected from hydroxy, halogen, alkoxy, alkyl, haloalkyl, amino, alkylamino, and cyano) fused to a WPTM2 ring;
WPTM5 is absent or an optionally substituted 5-6 membered aryl or heteroaryl ring (e.g., a 5-6 membered aryl or heteroaryl substituted with 0, 1, or 2 substituents selected from hydroxy, halogen, alkoxy, alkyl, haloalkyl, amino, alkylamino, and cyano);
WPTM6 and WPTM7 are independently optionally 4-7 cycloalkyl or heterocyclyl (e.g., 4-7 cycloalkyl or heterocyclyl each independently substituted with 0, 1, or 2 substituents selected from hydroxy, halogen, alkoxy, alkyl, haloalkyl, amino, alkylamino, and cyano), and the rings of WPTM6 and WPTM7 are fused or joined via a spiro linkage;
本明細書に記載される実施形態の態様のいずれにおいても、PTMは、式Iによって表されるか、

WPTM1は、任意に置換されたフェニルまたはピリジル(例えば、本明細書に記載されるように選択された追加の任意の置換基を有するかまたは有しないヒドロキシ置換基で置換されたフェニルなど、本明細書に記載されるように置換された)であり、
WPTM2は、任意に置換された6員ヘテロアリール環(例えば、アミノ基で置換されたピリダジンなど、本明細書に記載されるように置換された)であり、
WPTM3は、任意に置換された5~6員ヘテロアリール(例えば、ピラゾール、ピロール、イミダゾール、オキサゾール、オキサジアゾール、もしくはトリアゾール)、または任意に置換された4~9シクロアルキルもしくはヘテロシクリル環、任意に置換された架橋ビシクロアルキル、および架橋ビヘテロシクリル環が存在しない。
In any of the aspects of the embodiments described herein, the PTM is represented by formula I:
WPTM1 is an optionally substituted phenyl or pyridyl (e.g., substituted as described herein, such as phenyl substituted with a hydroxy substituent, with or without additional optional substituents selected as described herein);
WPTM2 is an optionally substituted 6-membered heteroaryl ring (e.g., substituted as described herein, such as pyridazine substituted with an amino group);
WPTM3 is free of optionally substituted 5-6 membered heteroaryl (e.g., pyrazole, pyrrole, imidazole, oxazole, oxadiazole, or triazole), or optionally substituted 4-9 cycloalkyl or heterocyclyl rings, optionally substituted bridged bicycloalkyl, and bridged biheterocyclyl rings.
本明細書に記載される態様または実施形態のいずれにおいても、PTMは、以下の式によって表されるか、
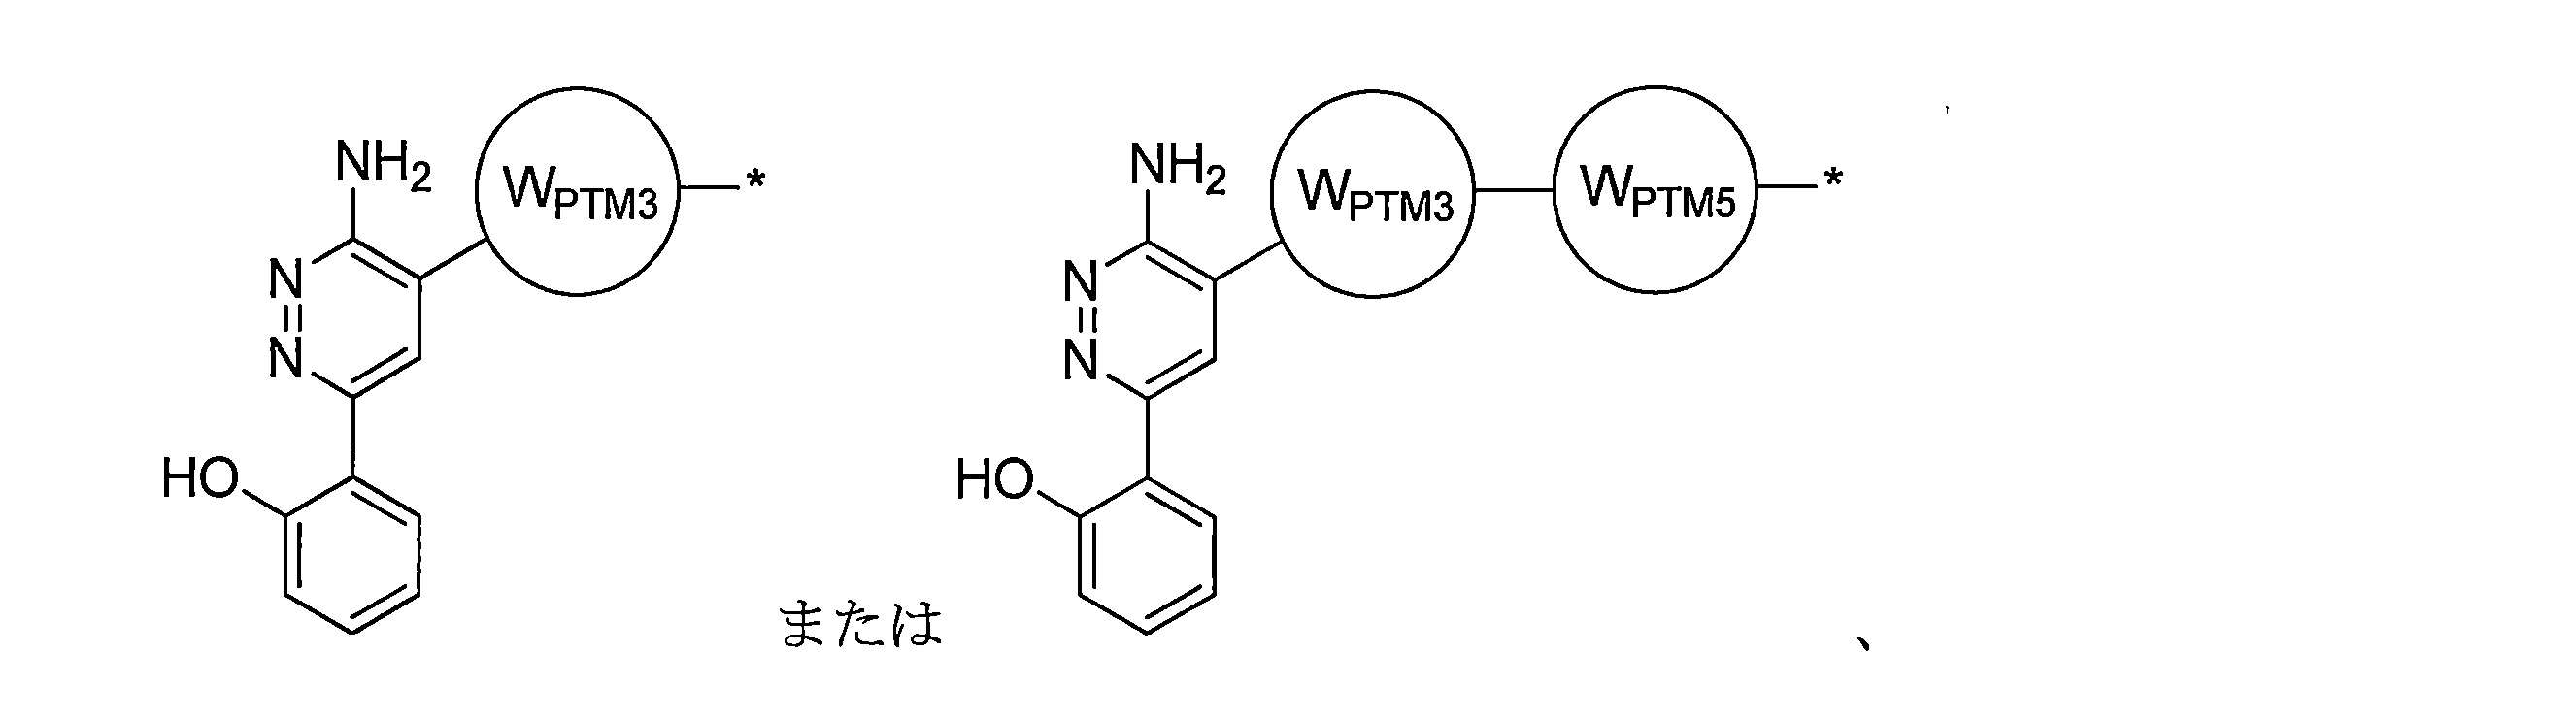
WPTM3は、不在であるか、または任意に置換された5~6員ヘテロアリール、任意に置換された4~9シクロアルキルもしくはヘテロシクリル環、任意に置換された架橋ビシクロアルキル、および架橋ビヘテロシクリル環であり、
WPTM5は、任意に置換された5~6員ヘテロアリールである。
In any of the aspects or embodiments described herein, the PTM may be represented by the formula:
WPTM3 is absent or an optionally substituted 5-6 membered heteroaryl, an optionally substituted 4-9 membered cycloalkyl or heterocyclyl ring, an optionally substituted bridged bicycloalkyl, and a bridged biheterocyclyl ring;
WPTM5 is an optionally substituted 5-6 membered heteroaryl.
本明細書に記載される態様または実施形態のいずれにおいても、PTMは、以下の式によって表されるか、
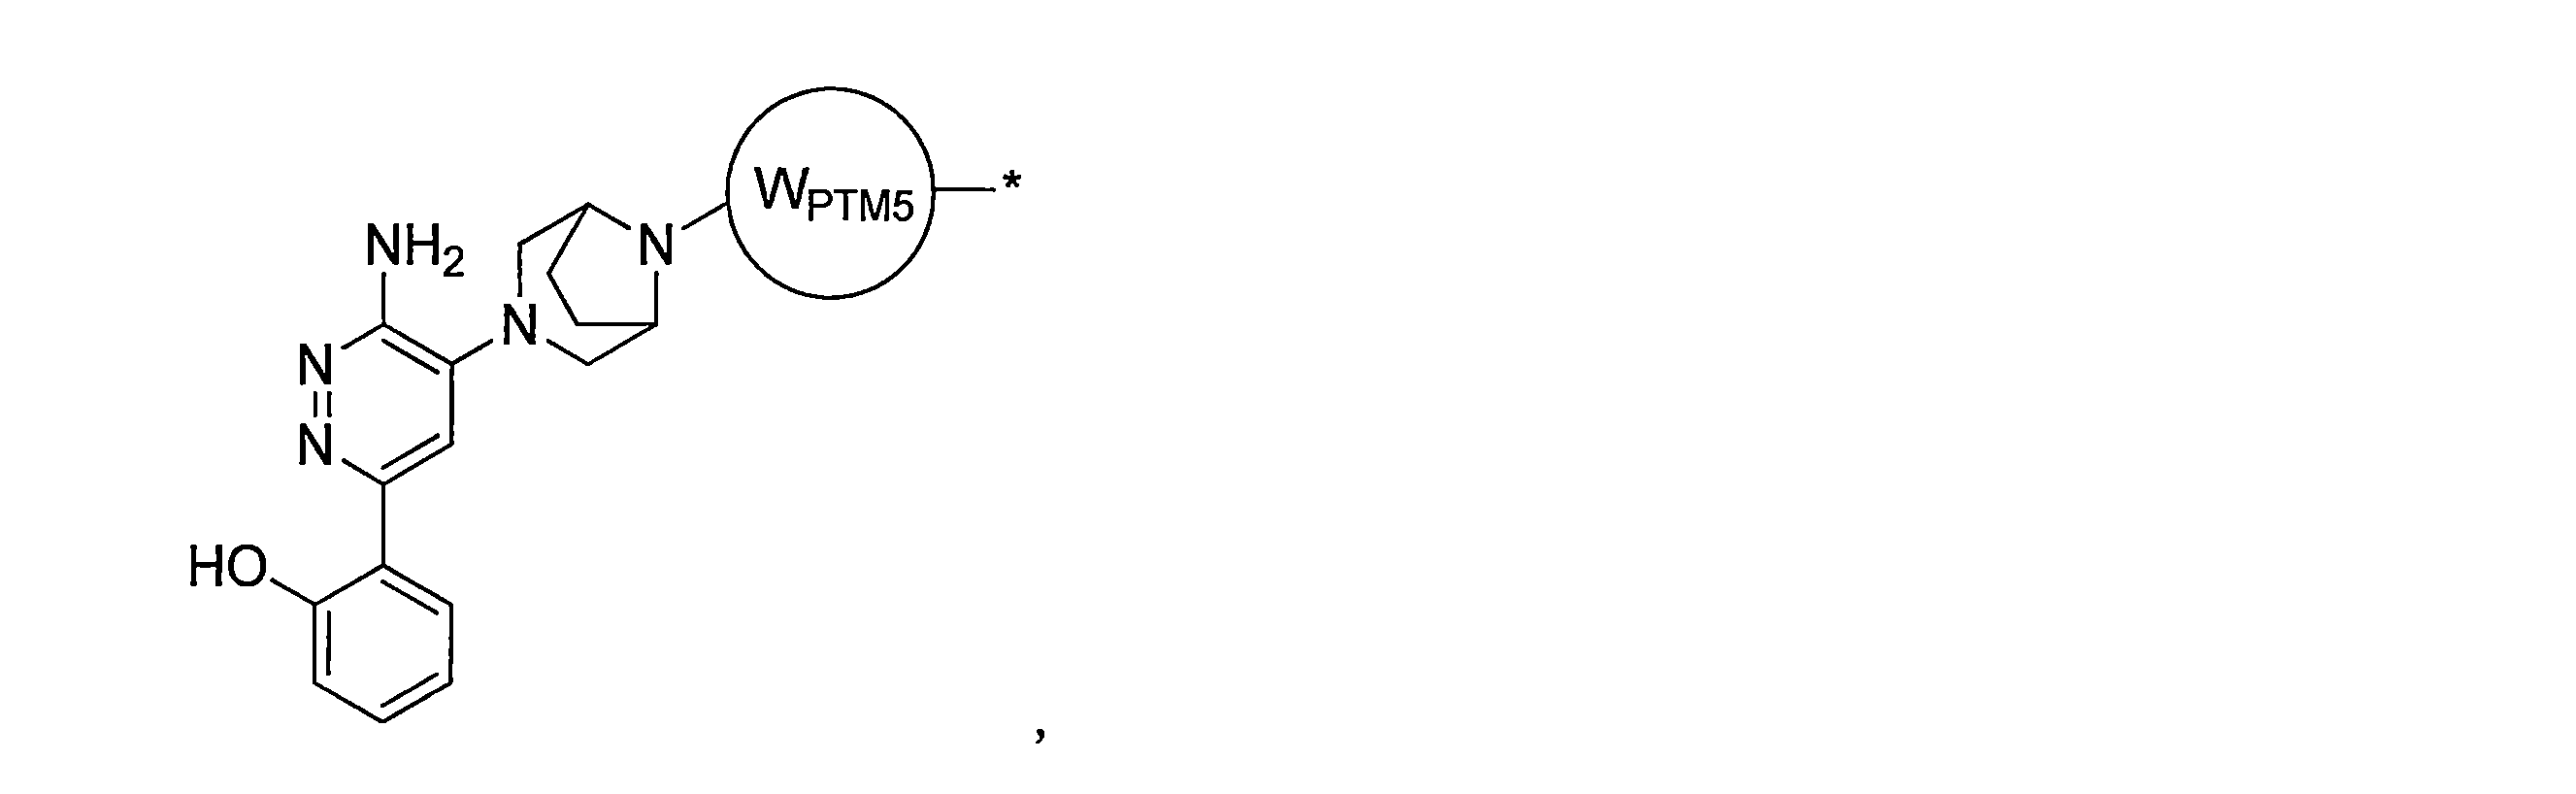
WPTM5は、フェニルまたはピリジニルである。
In any of the aspects or embodiments described herein, the PTM may be represented by the formula:
WPTM5 is phenyl or pyridinyl.
本明細書に記載される態様または実施形態のいずれにおいても、PTMは、以下の式によって表されるか、

本明細書に記載される態様または実施形態のいずれにおいても、PTMは、式IIIによって表されるか、
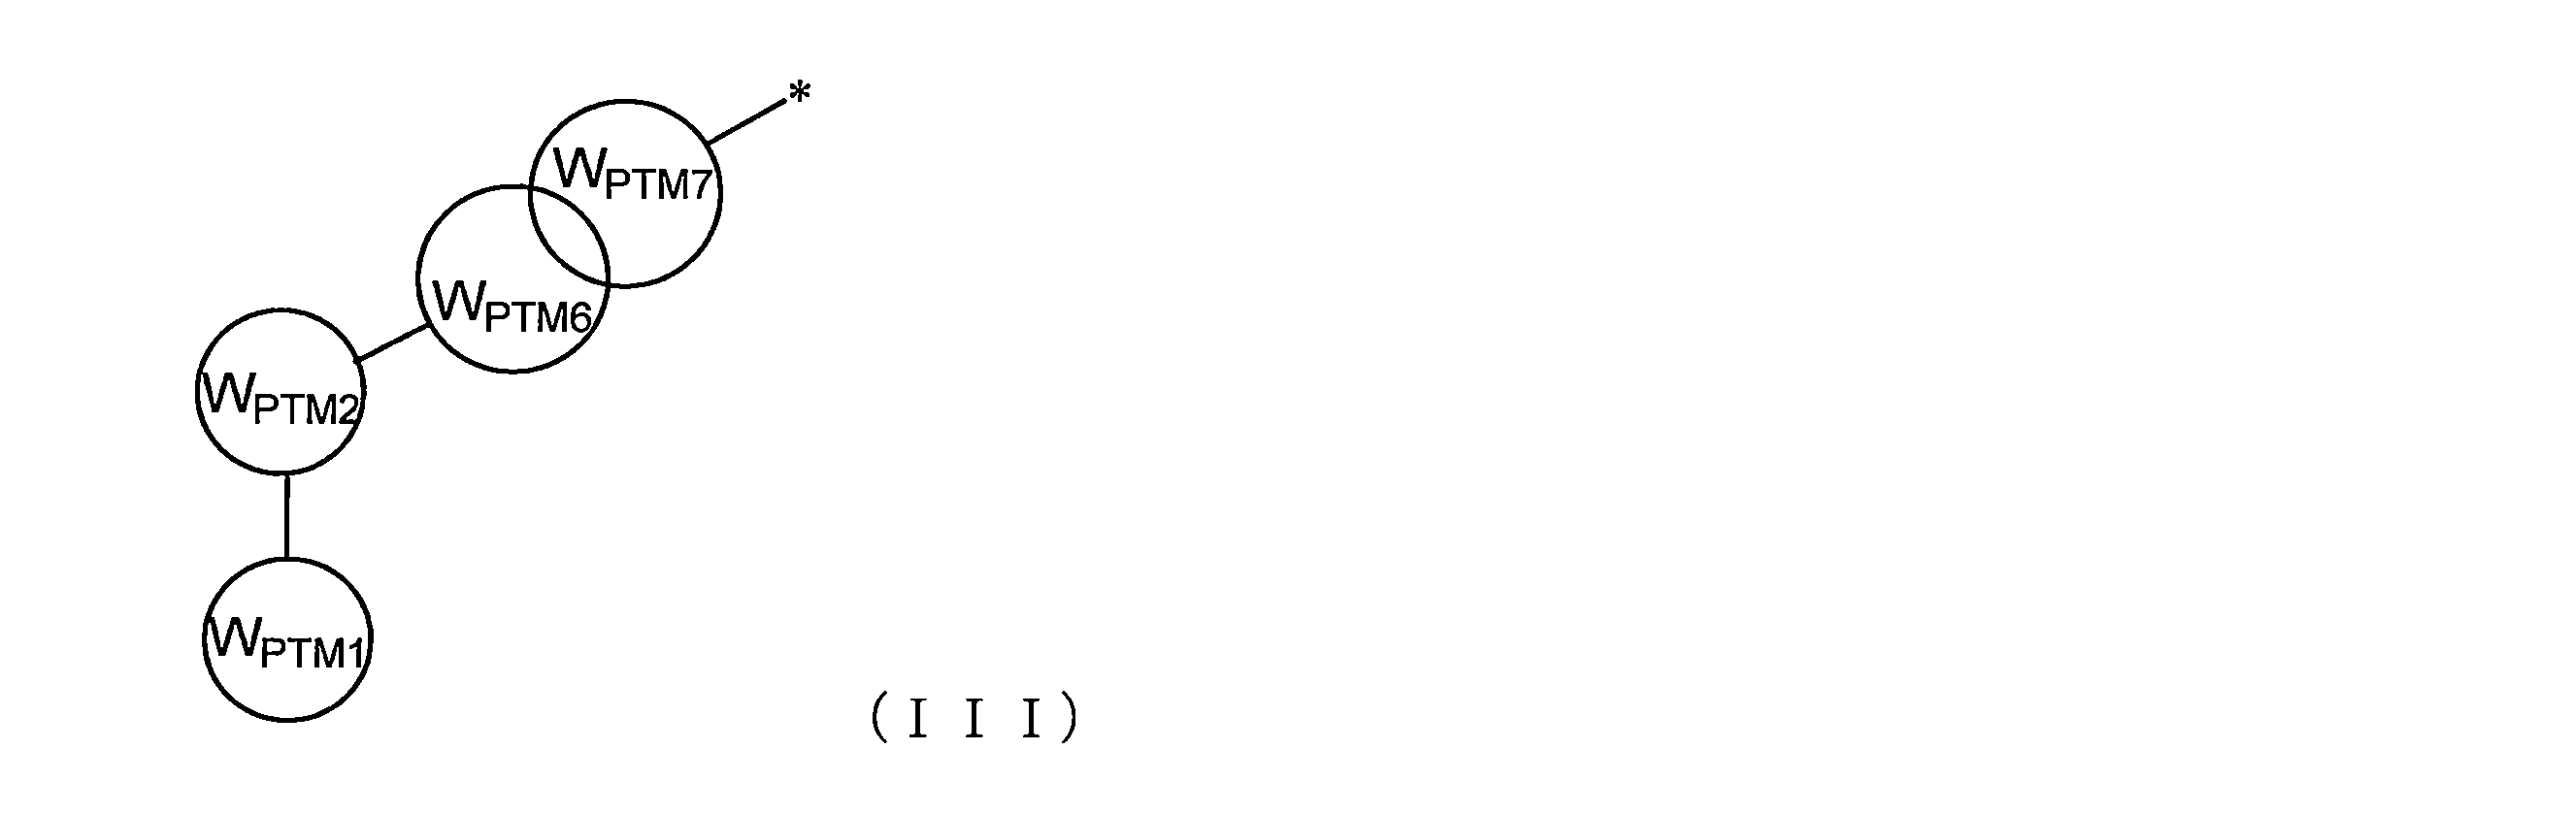
WPTM1は、追加の任意の置換基を有するかまたは有しないヒドロキシ置換基で置換されたフェニルであり、
WPTM2は、アミノ基で置換されたピリダジンであり、
WPTM6およびWPTM7は、スピロ環状環系(例えば、
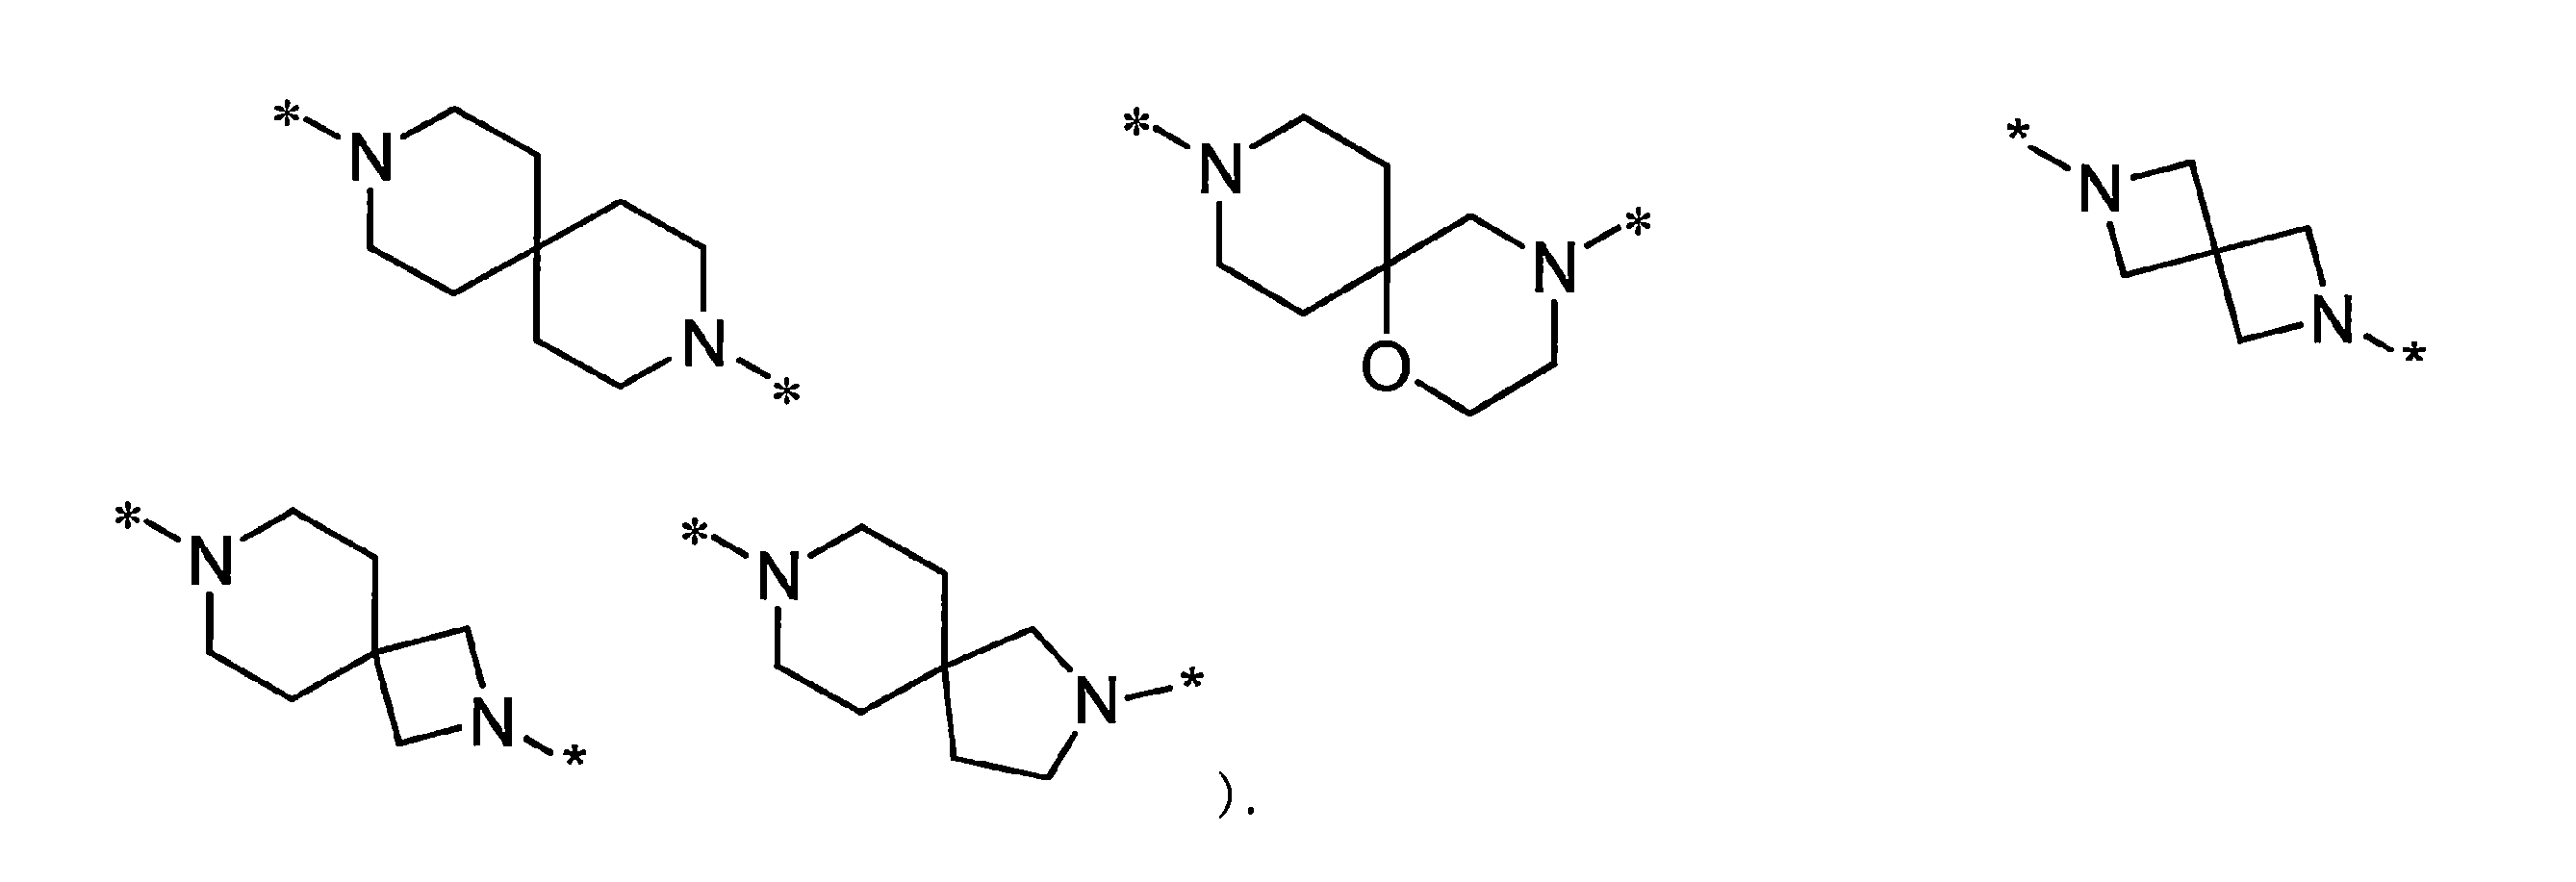
WPTM1 is a phenyl substituted with a hydroxy substituent, with or without additional optional substituents;
WPTM2 is an amino-substituted pyridazine;
WPTM6 and WPTM7 are spirocyclic ring systems, e.g.
本明細書に記載される態様または実施形態のいずれにおいても、PTMは、式IVaもしくはIVbによって表されるか、

WPTM1は、本明細書に記載される追加の任意の置換基を有するかまたは有しないヒドロキシ置換基で置換されたフェニルであり、
WPTM2は、アミノ基で置換されたピリダジンであり、
WPTM5は、存在しないか、またはピラゾール環もしくはピリジン環である。
In any of the aspects or embodiments described herein, the PTM is represented by formula IVa or IVb,
WPTM1 is phenyl substituted with a hydroxy substituent, with or without any additional substituents described herein;
WPTM2 is an amino-substituted pyridazine;
WPTM5 is absent or is a pyrazole ring or a pyridine ring.
本明細書に記載される態様または実施形態のいずれにおいても、PTMは、以下からなる群から選択される。

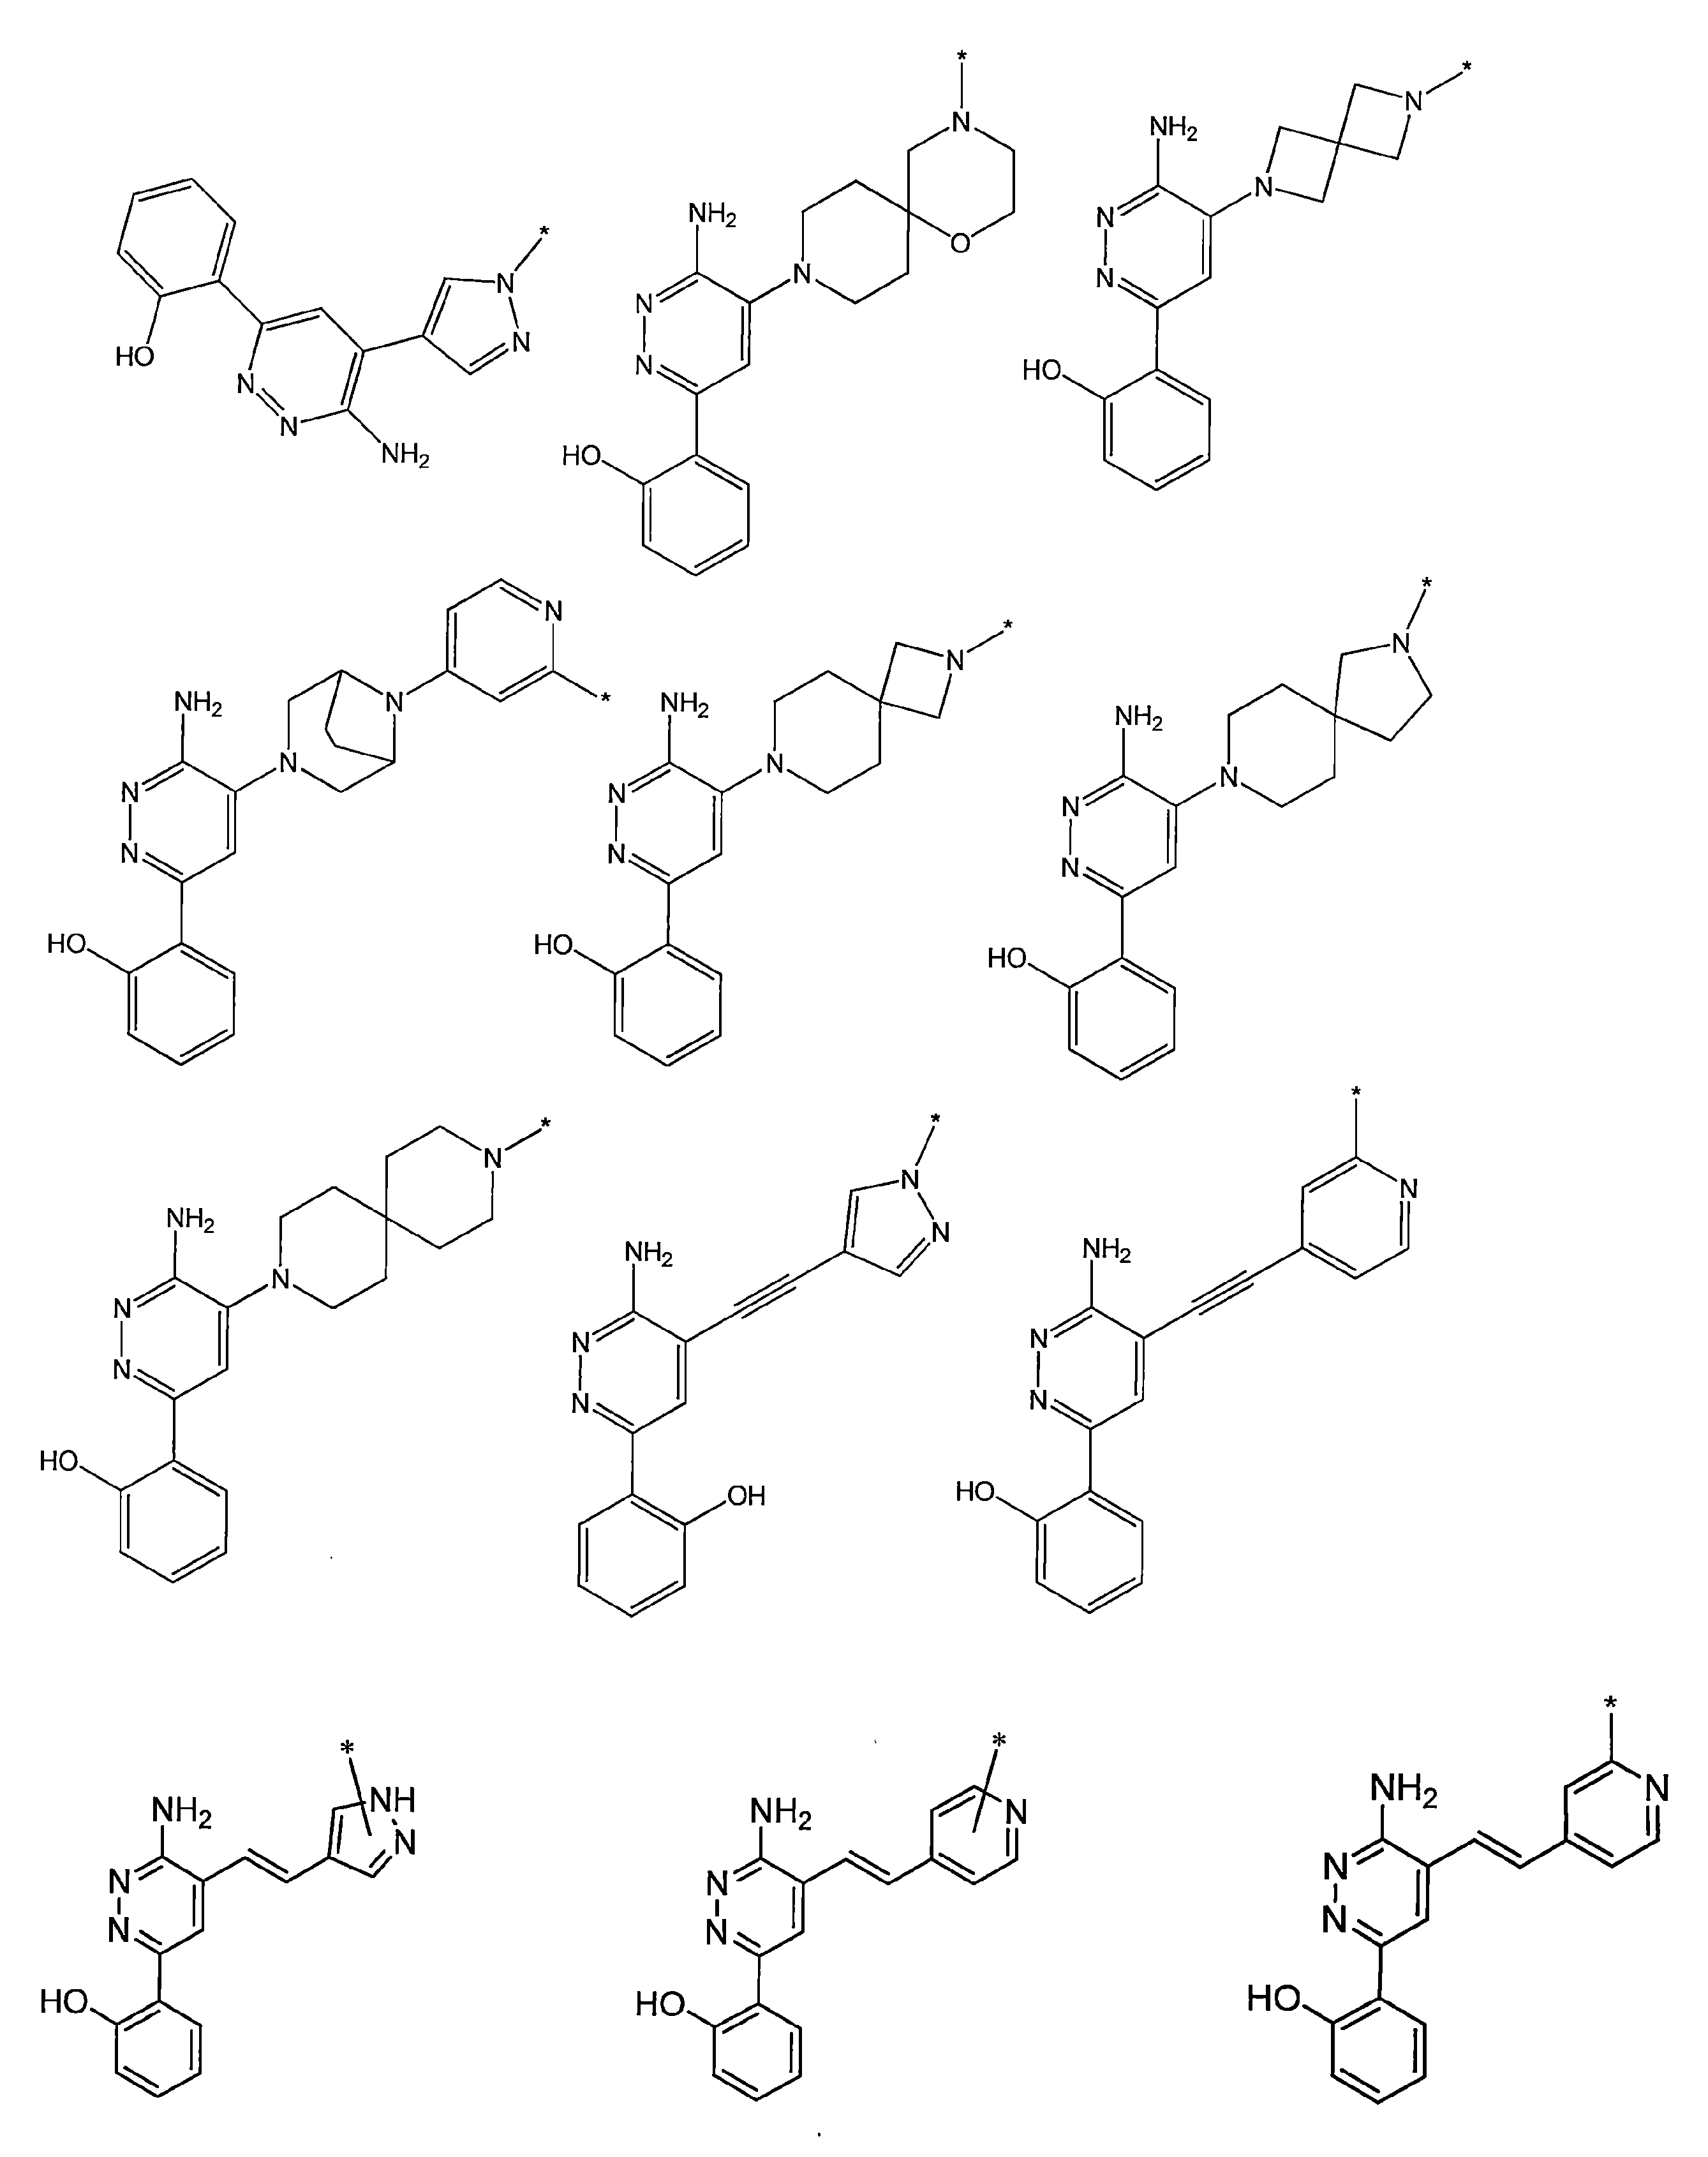

本明細書に記載される態様または実施形態のいずれにおいても、ULMは、以下によって表される化学構造であり、

W3は、任意に置換されたアリール、任意に置換されたヘテロアリール、または

R9およびR10は独立して、水素、任意に置換されたアルキル、任意に置換されたシクロアルキル、任意に置換されたヒドロキシアルキル、任意に置換されたヘテロアリール、またはハロアルキルであるか、またはR9、R10、およびそれらが結合する炭素原子が、任意に置換されたシクロアルキルを形成し、
R11は、任意に置換されたヘテロシクリル、任意に置換されたアルコキシ、任意に置換されたヘテロアリール、任意に置換されたアリール、

R12は、Hまたは任意に置換されたアルキルの群から選択され、
R13は、H、任意に置換されたアルキル、任意に置換されたアルキルカルボニル、任意に置換された(シクロアルキル)アルキルカルボニル、任意に置換されたアラルキルカルボニル、任意に置換されたアリールカルボニル、任意に置換された(ヘテロシクリル)カルボニル、または任意に置換されたアラルキルの群から選択され、
R14a、R14bは、各々独立して、H、ハロアルキル(例えば、フルオロアルキル)、任意に置換されたアルキル、任意に置換されたアルコキシ、任意に置換されたヒドロキシルアルキル、任意に置換されたアルキルアミン、任意に置換されたヘテロルキル、任意に置換されたアルキル-ヘテロシクロアルキル、任意に置換されたアルコキシ-ヘテロシクロアルキル、COR26、CONR27aR27b、NHCOR26、またはNHCH3COR26の群から選択され、R14aおよびR14bのうちの他方が、Hであり、またはR14a、R14bが、それらが結合している炭素原子と一緒になって、任意に置換された3~5員シクロアルキル、ヘテロシクロアルキル、スピロシクロアルキル、もしくはスピロヘテロシクリルを形成し、スピロヘテロシクリルが、エポキシドまたはアジリジンではなく、
W5は、任意に置換されたフェニル、任意に置換されたナプチル、または任意に置換された5~10員ヘテロアリールであり、
R15は、H、ハロゲン、CN、OH、NO2、NR27aR27b、OR27a、CONR27aR27b、NR27aCOR27b、SO2NR27aR27b、NR27aSO2R27b、任意に置換されたアルキル、任意に置換されたハロアルキル(例えば、任意に置換されたフルオロアルキル)、任意に置換されたハロアルコキシ、任意に置換されたアリール、任意に置換されたヘテロアリール、任意に置換されたシクロアルキル、または任意に置換されたヘテロシクリルの群から選択され、
各R16は、独立して、ハロ、CN、任意に置換されたアルキル、任意に置換されたハロアルキル、ヒドロキシ、または任意に置換されたハロアルコキシの群から選択され、
oは、0、1、2、3、または4であり、
R18は、H、ハロ、任意に置換されたアルコキシ、シアノ、任意に置換されたアルキル、ハロアルキル、ハロアルコキシまたはリンカーの群から独立して選択され、
各R26は、独立して、H、任意に置換されたアルキル、またはNR27aR27bから選択され、
各R27aおよびR27bは、独立して、H、任意に置換されたアルキル、任意に置換されたシクロアルキル(例えば、任意に置換された3~5員シクロアルキル)であるか、またはR27aおよびR27bは、それらが結合している窒素原子と一緒になって、4~6員ヘテロシクリルを形成し、
pは、0、1、2、3、または4であり、および式中、破線は、少なくとも1つのPTM、別のULM(ULM’)、または少なくとも1つのPTMもしくはULM’もしくはその両方をULMに結合する化学リンカー部分の結合部位を示す。
In any aspect or embodiment described herein, the ULM is a chemical structure represented by:
W3 is optionally substituted aryl, optionally substituted heteroaryl, or
R and R are independently hydrogen, optionally substituted alkyl, optionally substituted cycloalkyl, optionally substituted hydroxyalkyl, optionally substituted heteroaryl, or haloalkyl, or R, R and the carbon atom to which they are attached form an optionally substituted cycloalkyl;
R11 is optionally substituted heterocyclyl, optionally substituted alkoxy, optionally substituted heteroaryl, optionally substituted aryl,
R12 is selected from the group of H or optionally substituted alkyl;
R13 is selected from the group: H, optionally substituted alkyl, optionally substituted alkylcarbonyl, optionally substituted (cycloalkyl)alkylcarbonyl, optionally substituted aralkylcarbonyl, optionally substituted arylcarbonyl, optionally substituted (heterocyclyl)carbonyl, or optionally substituted aralkyl;
R14a, R14b are each independently selected from the group of H, haloalkyl (e.g., fluoroalkyl), optionally substituted alkyl, optionally substituted alkoxy, optionally substituted hydroxylalkyl, optionally substituted alkylamine, optionally substituted heteroalkyl, optionally substituted alkyl-heterocycloalkyl, optionally substituted alkoxy-heterocycloalkyl, COR26, CONR27aR27b, NHCOR26, or NHCH3COR26; the other of R14a and R14b is H; or R14a, R14b together with the carbon atom to which they are attached form an optionally substituted 3-5 membered cycloalkyl, heterocycloalkyl, spirocycloalkyl, or spiroheterocyclyl, wherein the spiroheterocyclyl is not an epoxide or aziridine;
W5 is an optionally substituted phenyl, an optionally substituted naphthyl, or an optionally substituted 5-10 membered heteroaryl;
R15 is selected from the group of H, halogen, CN, OH, NO2, NR27aR27b, OR27a, CONR27aR27b, NR27aCOR27b, SO2NR27aR27b, NR27aSO2R27b, optionally substituted alkyl, optionally substituted haloalkyl (e.g., optionally substituted fluoroalkyl), optionally substituted haloalkoxy, optionally substituted aryl, optionally substituted heteroaryl, optionally substituted cycloalkyl, or optionally substituted heterocyclyl;
Each R16 is independently selected from the group of halo, CN, optionally substituted alkyl, optionally substituted haloalkyl, hydroxy, or optionally substituted haloalkoxy;
o is 0, 1, 2, 3, or 4;
R18 is independently selected from the group of H, halo, optionally substituted alkoxy, cyano, optionally substituted alkyl, haloalkyl, haloalkoxy, or a linker;
Each R26 is independently selected from H, optionally substituted alkyl, or NR27aR27b;
each R27a and R27b is independently H, optionally substituted alkyl, optionally substituted cycloalkyl (e.g., optionally substituted 3- to 5-membered cycloalkyl), or R27a and R27b together with the nitrogen atom to which they are attached form a 4- to 6-membered heterocyclyl;
p is 0, 1, 2, 3, or 4, and wherein the dashed lines indicate the site of attachment of at least one PTM, another ULM (ULM'), or a chemical linker moiety linking at least one PTM or ULM' or both to a ULM.
本明細書に記載される態様または実施形態のいずれにおいても、ULMは、以下の群から選択される化学構造を有し、

R1は、H、任意に置換されたアルキル、任意に置換されたシクロアルキル、任意に置換されたヒドロキシアルキル、任意に置換されたヘテロアリール、またはハロアルキルであり、
R14aは、H、ハロアルキル、任意に置換されたアルキル、メチル、フルオロメチル、ヒドロキシメチル、エチル、イソプロピル、またはシクロプロピルであり、
R15は、H、ハロゲン、CN、OH、NO2、任意に置換されたヘテロアリール、任意に置換されたアリール、任意に置換されたアルキル(例えば、任意に置換されたハロアルキル)、任意に置換されたハロアルキル、任意に置換されたハロアルコキシ、任意に置換されたシクロアルキル、または任意に置換されたヘテロシクリルからなる群から選択され、
Xは、C、CH2、またはC=Oであり、
R3は、存在しないか、または任意に置換された5もしくは6員ヘテロアリールであり、
破線は、少なくとも1つのPTM、別のULM(ULM’)、または少なくとも1つのPTMもしくはULM’もしくはその両方をULMに結合する化学リンカー部分の結合部位を示す。
In any aspect or embodiment described herein, the ULM has a chemical structure selected from the group consisting of:
R1 is H, optionally substituted alkyl, optionally substituted cycloalkyl, optionally substituted hydroxyalkyl, optionally substituted heteroaryl, or haloalkyl;
R14a is H, haloalkyl, optionally substituted alkyl, methyl, fluoromethyl, hydroxymethyl, ethyl, isopropyl, or cyclopropyl;
R15 is selected from the group consisting of H, halogen, CN, OH, NO2, optionally substituted heteroaryl, optionally substituted aryl, optionally substituted alkyl (e.g., optionally substituted haloalkyl), optionally substituted haloalkyl, optionally substituted haloalkoxy, optionally substituted cycloalkyl, or optionally substituted heterocyclyl;
X is C, CH2, or C=O;
R3 is absent or is an optionally substituted 5- or 6-membered heteroaryl;
The dashed lines indicate the attachment sites of at least one PTM, another ULM (ULM'), or a chemical linker moiety linking at least one PTM or ULM', or both, to a ULM.
本明細書に記載される態様または実施形態のいずれにおいても、ULMは、以下の化学構造に従う基、

ULM-gのR1’は、任意に置換されたC1-C6アルキル基、任意に置換された-(CH2)nOH、任意に置換された-(CH2)nSH、任意に置換された(CH2)n-O-(C1-C6)アルキル基、エポキシド部分WCOCW(式中、各Wは、独立して、HまたはC1-C3アルキル基である)を含有する任意に置換された(CH2)n-WCOCW-(C0-C6)アルキル基、任意に置換された-(CH2)nCOOH、任意に置換された-(CH2)nC(O)-(C1-C6アルキル)、任意に置換された-(CH2)nNHC(O)-R″、任意に置換された-(CH2)nC(O)-N(R″)2、任意に置換された-(CH2)nOC(O)-N(R″)2、-(CH2O)nH、任意に置換された-(CH2)nOC(O)-(C1-C6アルキル)、任意に置換された-(CH2)nC(O)-O-(C1-C6アルキル)、任意に置換された-(CH2O)nCOOH、任意に置換された-(OCH2)nO-(C1-C6アルキル)、任意に置換された-(CH2O)nC(O)-(C1-C6アルキル)、任意に置換された-(OCH2)nNHC(O)-R″、任意に置換された-(CH2O)nC(O)-N(R″)2、-(CH2CH2O)nH、任意に置換された-(CH2CH2O)nCOOH、任意に置換された-(OCH2CH2)nO-(C1-C6アルキル)、任意に置換された-(CH2CH2O)nC(O)-(C1-C6アルキル)、任意に置換された-(OCH2CH2)nNHC(O)-R″、任意に置換された-(CH2CH2O)nC(O)-N(R″)2、任意に置換された-SO2RS、任意に置換されたS(O)RS、NO2、CN、またはハロゲン(F、Cl、Br、I、好ましくは、FまたはCl)であり、
ULM-gの各R″は、独立して、H、または1個もしくは2個のヒドロキシル基または最大3個のハロゲン基(好ましくは、フッ素)で任意に置換され得るC1-C6アルキル基であり、
ULM-gのRSは、C1-C6アルキル基、任意に置換されたアリール、ヘテロアリール、もしくはヘテロシクリル基、または-(CH2)mN(R″)2基であり、
ULM-gのXおよびX’は、各々独立して、C=O、C=S、-S(O)、S(O)2であり(好ましくは、XおよびX’は、両方ともC=Oであり)、
ULM-gのR2’は、任意に置換された-(CH2)n-(C=O)u(NR″)v(SO2)wアルキリ基、任意に置換された-(CH2)n-(C=O)u(NR″)v(SO2)wNR1NR2N基、任意に置換された-(CH2)n-(C=O)u(NR″)v(SO2)w-アリール、任意に置換された-(CH2)n-(C=O)u(NR″)v(SO2)w-ヘテロアリール、任意に置換された-(CH2)n-(C=O)vNR″(SO2)w-ヘテロシクリル、任意に置換された-NR″-(CH2)n-C(O)u(NR″)v(SO2)w-アルキル、任意に置換された-NR″-(CH2)n-C(O)u(NR″)v(SO2)w-NR1NR2N、任意に置換された-NR″-(CH2)n-C(O)u(NR″)v(SO2)w-NR″C(O)R1N、任意に置換された-NR″-(CH2)n-(C=O)u(NR″)v(SO2)w-アリール、任意に置換された-NR″-(CH2)n-(C=O)u(NR″)v(SO2)w-ヘテロアリール、もしくは任意に置換された-NR″-(CH2)n-(C=O)vNR″(SO2)w-ヘテロシクリル、任意に置換された-XR2’-アルキリ基、任意に置換された-XR2’-アリール基、任意に置換された-XR2’-ヘテロアリール基、任意に置換された-XR2’-ヘテロシクリル基であり、
ULM-gのR3’は、任意に置換されたアルキル、任意に置換された-(CH2)n-(O)u(NR″)v(SO2)w-アルキル、任意に置換された-(CH2)n-C(O)u(NR″)v(SO2)w-NR1NR2N、任意に置換された-(CH2)n-C(O)u(NR″)v(SO2)w-NR″C(O)R1N、任意に置換された-(CH2)n-C(O)u(NR″)v(SO2)w-C(O)(R″)2、任意に置換された-(CH2)n-C(O)u(NR″)v(SO2)w-アリール、任意に置換された-(CH2)n-C(O)u(NR″)v(SO2)w-ヘテロアリール、任意に置換された-(CH2)n-C(O)u(NR″)v(SO2)w-ヘテロシクリル、任意に置換された-NR″-(CH2)n-C(O)u(NR″)v(SO2)w-アルキル、任意に置換された-NR″-(CH2)n-C(O)u(NR″)v(SO2)w-NR1NR2N、任意に置換された-NR″-(CH2)n-C(O)u(NR″)v(SO2)w-NR″C(O)R1N、任意に置換された-NR″-(CH2)n-C(O)u(NR″)v(SO2)w-アリール、任意に置換された-NR″-(CH2)n-C(O)u(NR″)v(SO2)w-ヘテロアリール、任意に置換された-NR1-(CH2)n-C(O)u(NR″)v(SO2)w-ヘテロシクリル、任意に置換された-O-(CH2)n-(C=O)u(NR″)v(SO2)w-アルキル、任意に置換された-O-(CH2)n-(C=O)u(NR″)v(SO2)w-NR1NR2N、任意に置換された-O-(CH2)n-(C=O)u(NR″)v(SO2)w-NR″C(O)R1N、任意に置換された-O-(CH2)n-(C=O)u(NR″)v(SO2)w-アリール、任意に置換された-O-(CH2)n-(C=O)u(NR″)v(SO2)w-ヘテロアリール、もしくは任意に置換された-O-(CH2)n-(C=O)u(NR″)v(SO2)w-ヘテロシクリル、-(CH2)n-(V)n’-(CH2)n-(V)n’-アルキル基、任意に置換された-(CH2)n-(V)n’-(CH2)n-(V)n’-アリール基、任意に置換された-(CH2)n-(V)n’-(CH2)n-(V)n’-ヘテロアリール基、任意に置換された-(CH2)n-(V)n’-(CH2)n-(V)n’-ヘテロシクリル’基、任意に置換された-(CH2)n-N(R1’)(C=O)m’-(V)n’-アルキル基、任意に置換された-(CH2)n-N(R1’)(C=O)m’-(V)n’-アリール基、任意に置換された-(CH2)n-N(R1’)(C=O)m’-(V)n’-ヘテロアリール基、任意に置換された-(CH2)n-N(R1’)(C=O)m’-(V)n’-ヘテロシクリル基、任意に置換された-XR3’-アルキル基、任意に置換された-XR3’-アリール基、任意に置換された-XR3’-ヘテロアリール基、任意に置換された-XR3’-ヘテロシクリル基であり、
ULM-gのR1NおよびR2Nは、各々独立して、H、1個もしくは2個のヒドロキシル基および最大3個のハロゲン基で任意に置換されたC1-C6アルキル、または任意に置換された-(CH2)n-アリール、-(CH2)n-ヘテロアリール、もしくは-(CH2)n-ヘテロシクリル基であり、
ULM-gのVは、O、SまたはNR1であり、
ULM-gの各R1’は、独立して、H、またはC1-C3アルキル基であり、
ULM-gのXR2’およびXR3’は、各々独立して、任意に置換された-CH2)n-、-CH2)n-CH(Xv)=CH(Xv)-(cisまたはtrans)、-CH2)n-CH≡CH-、-(CH2CH2O)n-、またはC3-C6シクロアルキル基であり、式中、Xvは、H、ハロ、または任意に置換されたC1-C3アルキル基であり、
ULM-gの各mは独立して、0、1、2、3、4、5、6であり、
ULM-gの各m’は独立して、0または1であり、
ULM-gの各nは独立して、0、1、2、3、4、5、6であり、
ULM-gの各n’は独立して、0または1であり、
ULM-gの各uは独立して、0または1であり、
ULM-gの各vは独立して、0または1であり、
ULM-gの各wは独立して、0または1であり、および
PTMがULM’でない場合、ULM-gのR1’、R2’、R3’、X、およびX’のうちのいずれか1つ以上が任意に修飾されて、リンカー基を介してPTM基に共有結合するか、またはPTMがULM’である場合、ULMおよびULM’の各々のR1’、R2’、R3’、X、およびX’のうちのいずれか1つ以上は任意に修飾されて、直接もしくはそのリンカー基を介して互いに共有結合する。
In any of the aspects or embodiments described herein, the ULM may be a group according to the following chemical structure:
R1' of ULM-g is an optionally substituted C1-C6 alkyl group, an optionally substituted -(CH2)nOH, an optionally substituted -(CH2)nSH, an optionally substituted (CH2)n-O-(C1-C6)alkyl group, an optionally substituted (CH2)n-WCOCW-(C0-C6)alkyl group containing an epoxide moiety WCOCW (where each W is independently H or a C1-C3 alkyl group), an optionally substituted -(CH 2) nCOOH, optionally substituted -(CH2)nC(O)-(C1-C6 alkyl), optionally substituted -(CH2)nNHC(O)-R", optionally substituted -(CH2)nC(O)-N(R")2, optionally substituted -(CH2)nOC(O)-N(R")2, -(CH2O)nH, optionally substituted -(CH2)nOC(O)-(C1-C6 alkyl), optionally substituted -(CH2)nC(O)-O-(C1-C6 optionally substituted -(CH2O)nCOOH, optionally substituted -(OCH2)nO-(C1-C6 alkyl), optionally substituted -(CHO)nC(O)-(C1-C6 alkyl), optionally substituted -(OCH2)nNHC(O)-R", optionally substituted -(CHO)nC(O)-N(R")2, -(CH2CH2O)nH, optionally substituted -(CH2CH2O)nCOOH, optionally substituted -( optionally substituted —(CHCHO)C(O)—N(R″)2, optionally substituted —SORS, optionally substituted S(O)RS, NO, CN, or halogen (F, Cl, Br, I, preferably F or Cl);
Each R″ in ULM-g is independently H or a C1-C6 alkyl group which may be optionally substituted with one or two hydroxyl groups or up to three halogen groups (preferably fluorine);
RS of ULM-g is a C1-C6 alkyl group, an optionally substituted aryl, heteroaryl, or heterocyclyl group, or a -(CH2)mN(R'')2 group;
X and X' of ULM-g are each independently C=O, C=S, -S(O), S(O)2 (preferably, X and X' are both C=O);
R2' of ULM-g is an optionally substituted -(CH2)n-(C=O)u(NR'')v(SO2)w alkyl group, an optionally substituted -(CH2)n-(C=O)u(NR'')v(SO2)w NR1NR2N group, an optionally substituted -(CH2)n-(C=O)u(NR'')v(SO2)w aryl, an optionally substituted -(CH2)n-(C=O)u(NR'')v(SO2)w-heteroaryl, an optionally substituted -(CH2)n-(C=O)vNR''(SO2)w-heterocyclyl, an optionally substituted -NR''-(CH2)n-C(O)u(NR'')v(SO2)w alkyl, an optionally substituted -NR''-(CH2)n-C(O)u(NR'')v (SO2)w-NR1NR2N, optionally substituted -NR"-(CH2)n-C(O)u(NR")v(SO2)w-NR"C(O)RN, optionally substituted -NR"-(CH2)n-(C=O)u(NR")v(SO2)w-aryl, optionally substituted -NR"-(CH2)n-(C=O)u(NR")v(SO2)w-heteroaryl or optionally substituted -NR"-(CH2)n-(C=O)vNR"(SO2)w-heterocyclyl, optionally substituted -XR2'-alkyl group, optionally substituted -XR2'-aryl group, optionally substituted -XR2'-heteroaryl group, or optionally substituted -XR2'-heterocyclyl group;
R3' of ULM-g is optionally substituted alkyl, optionally substituted -(CH2)n-(O)u(NR'')v(SO2)w-alkyl, optionally substituted -(CH2)n-C(O)u(NR'')v(SO2)w-NR1NR2N, optionally substituted -(CH2)n-C(O)u(NR'')v(SO2)w-NR''C(O)R1N, optionally substituted -(C H2)n-C(O)u(NR'')v(SO2)w-C(O)(R'')2, optionally substituted -(CH2)n-C(O)u(NR'')v(SO2)w-aryl, optionally substituted -(CH2)n-C(O)u(NR'')v(SO2)w-heteroaryl, optionally substituted -(CH2)n-C(O)u(NR'')v(SO2)w-heterocyclyl, optionally substituted optionally substituted -NR''-(CH2)n-C(O)u(NR'')v(SO2)w-alkyl, optionally substituted -NR''-(CH2)n-C(O)u(NR'')v(SO2)w-NR1NR2N, optionally substituted -NR''-(CH2)n-C(O)u(NR'')v(SO2)w-NR''C(O)R1N, optionally substituted -NR''-(CH2)n-C(O)u(NR'')v(SO2)w-NR''C(O)R1N optionally substituted —NR″-(CH2)n-C(O)u(NR″)v(SO2)w-aryl, optionally substituted —NR″-(CH2)n-C(O)u(NR″)v(SO2)w-heteroaryl, optionally substituted —NR1-(CH2)n-C(O)u(NR″)v(SO2)w-heterocyclyl, optionally substituted —O—(CH2)n-(C═O)u(NR″)v(SO2)w-alkyl, optionally substituted —O—( CH2)n-(C=O)u(NR'')v(SO2)w-NR1NR2N, optionally substituted -O-(CH2)n-(C=O)u(NR'')v(SO2)w-NR''C(O)R1N, optionally substituted -O-(CH2)n-(C=O)u(NR'')v(SO2)w-aryl, optionally substituted -O-(CH2)n-(C=O)u(NR'')v(SO2)w- Heteroaryl, or optionally substituted -O-(CH2)n-(C=O)u(NR'')v(SO2)w-heterocyclyl, -(CH2)n-(V)n'-(CH2)n-(V)n'-alkyl group, optionally substituted -(CH2)n-(V)n'-(CH2)n-(V)n'-aryl group, optionally substituted -(CH2)n-(V)n'-(CH2)n-( V) n'-heteroaryl groups, optionally substituted -(CH2)n-(V) n'-(CH2)n-(V) n'-heterocyclyl' groups, optionally substituted -(CH2)n-N(R1')(C=O)m'-(V) n'-alkyl groups, optionally substituted -(CH2)n-N(R1')(C=O)m'-(V) n'-aryl groups, optionally substituted -(CH2)n-N(R1')(C=O)m'-(V) n'-heteroaryl groups, optionally substituted -(CH2)n-N(R1')(C=O)m'-(V) n'-heterocyclyl groups, optionally substituted -XR3'-alkyl groups, optionally substituted -XR3'-aryl groups, optionally substituted -XR3'-heteroaryl groups, optionally substituted -XR3'-heterocyclyl groups;
R1N and R2N of ULM-g are each independently H, C1-C6 alkyl optionally substituted with one or two hydroxyl groups and up to three halogen groups, or an optionally substituted -(CH2)n-aryl, -(CH2)n-heteroaryl, or -(CH2)n-heterocyclyl group;
V of ULM-g is O, S or NR1;
Each R1' in ULM-g is independently H or a C1-C3 alkyl group;
XR2' and XR3' of ULM-g are each independently an optionally substituted -CH2)n-, -CH2)n-CH(Xv)=CH(Xv)-(cis or trans), -CH2)n-CH≡CH-, -(CH2CHO)n-, or a C3-C6 cycloalkyl group, where Xv is H, halo, or an optionally substituted C1-C3 alkyl group;
Each m in ULM-g is independently 0, 1, 2, 3, 4, 5, or 6;
Each m' in ULM-g is independently 0 or 1;
Each n in ULM-g is independently 0, 1, 2, 3, 4, 5, or 6;
Each n' in ULM-g is independently 0 or 1;
Each u in ULM-g is independently 0 or 1;
Each v in ULM-g is independently 0 or 1;
Each w of ULM-g is independently 0 or 1; and when the PTM is not a ULM', any one or more of R1', R2', R3', X, and X' of ULM-g are optionally modified to be covalently bonded to the PTM group via a linker group, or when the PTM is a ULM', any one or more of R1', R2', R3', X, and X' of each of the ULM and ULM' are optionally modified to be covalently bonded to each other directly or via their linker groups.
本明細書に記載される態様または実施形態のいずれにおいても、ULMは、以下の式:
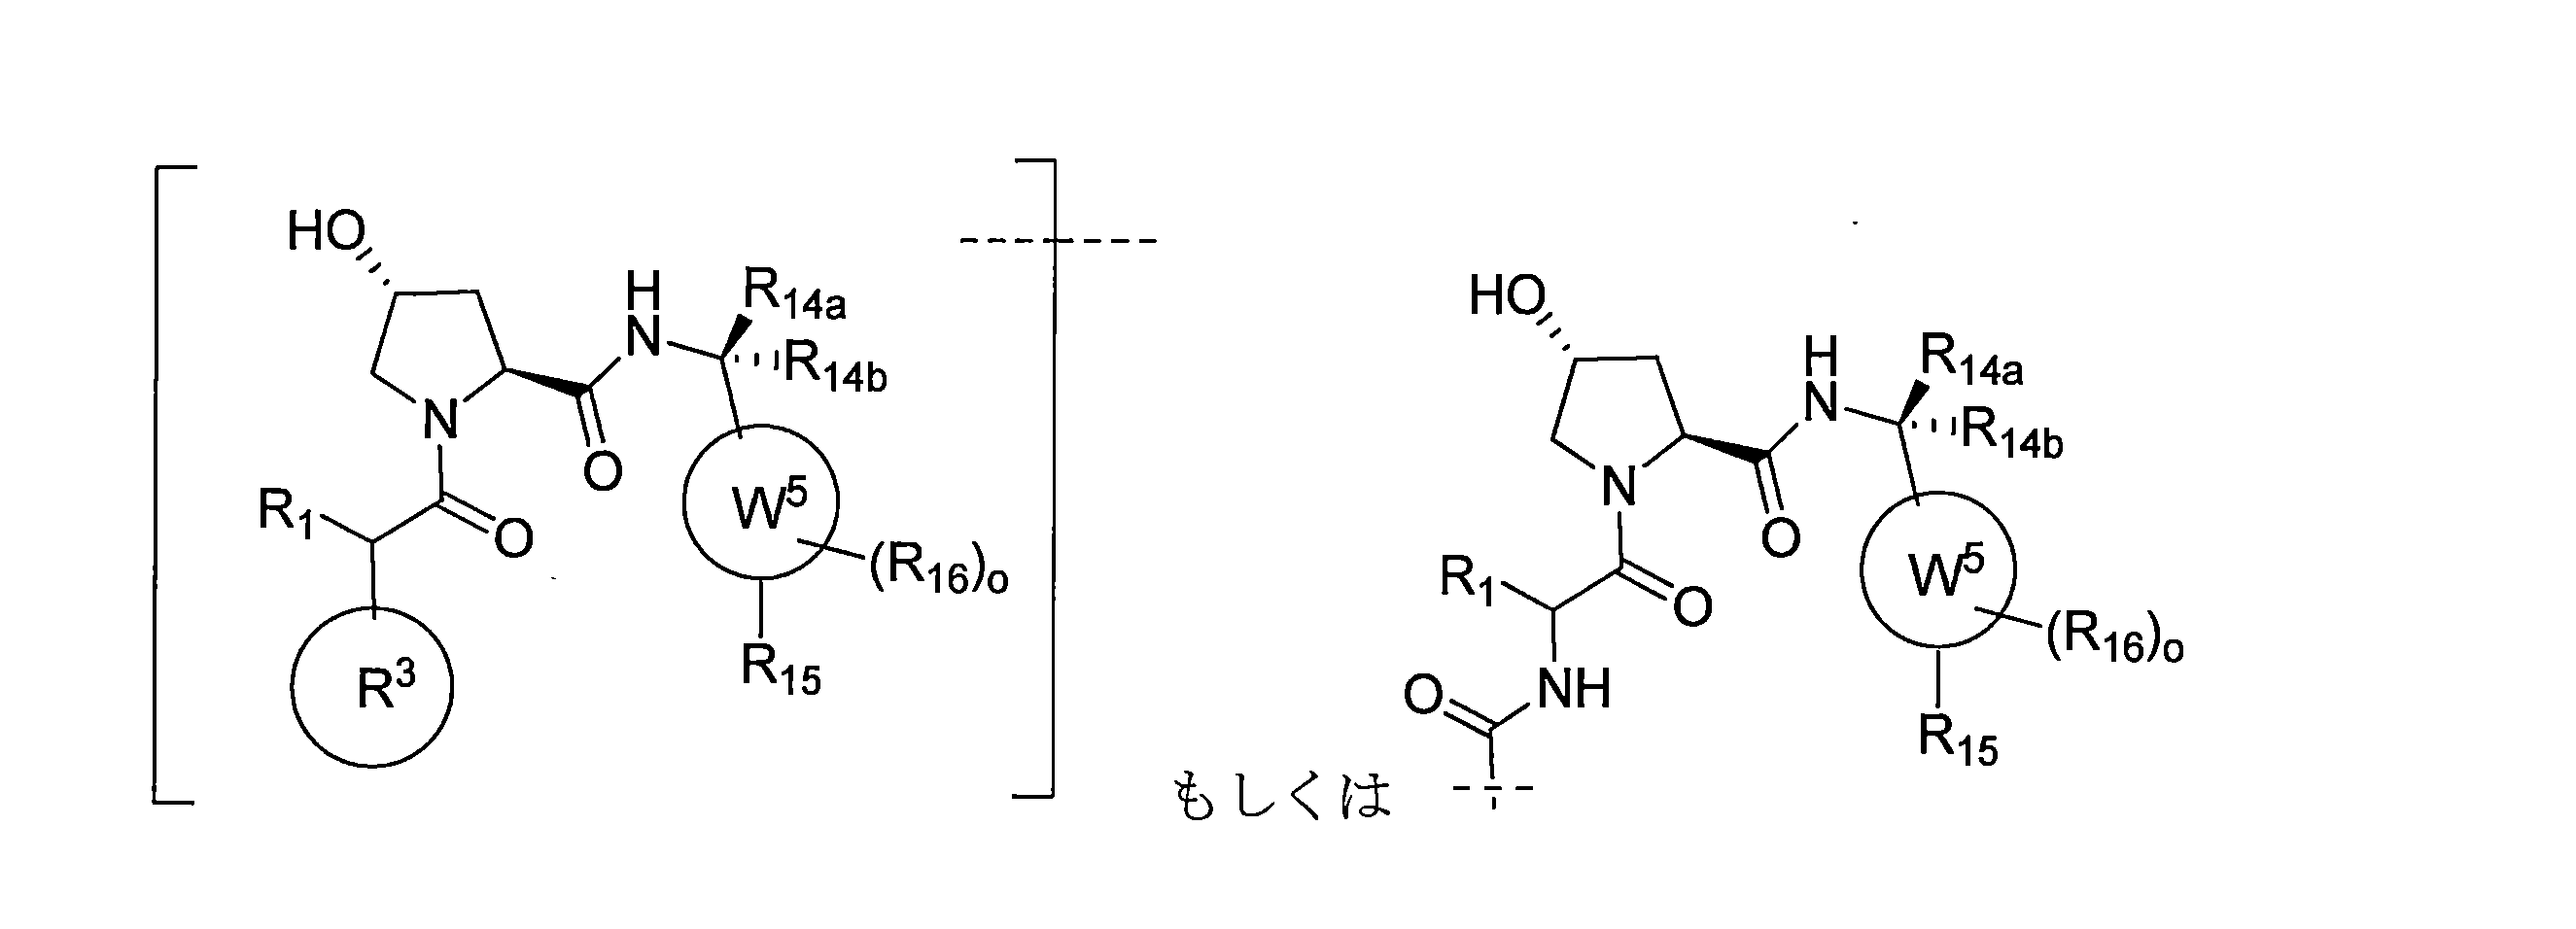
R1は、H、任意に置換されたアルキル、または任意に置換されたシクロアルキルであり、
R3は、任意に置換された5~6員ヘテロアリールであり、
W5は、任意に置換されたフェニル、任意に置換されたナプチル、または任意に置換されたピリジニルであり、
R14aおよびR14bのうちの一方は、H、任意に置換されたアルキル、任意に置換されたハロアルキル、任意に置換されたアルコキシ、任意に置換されたヒドロキシルアルキル、任意に置換されたアルキルアミン、任意に置換されたヘテロルキル、任意に置換されたアルキル-ヘテロシクロアルキル、任意に置換されたアルコキシ-ヘテロシクロアルキル、COR26、CONR27aR27b、NHCOR26、もしくはNHCH3COR26であり、R14aおよびR14bのうちの他方は、Hであり、またはR14a、R14bは、それらが結合している炭素原子と一緒になって、任意に置換された3~5員シクロアルキル、ヘテロシクロアルキル、スピロシクロアルキル、もしくはスピロヘテロシクリルを形成し、スピロヘテロシクリルが、エポキシドまたはアジリジンではなく、
R15は、CN、任意に置換されたフルオロアルキル、


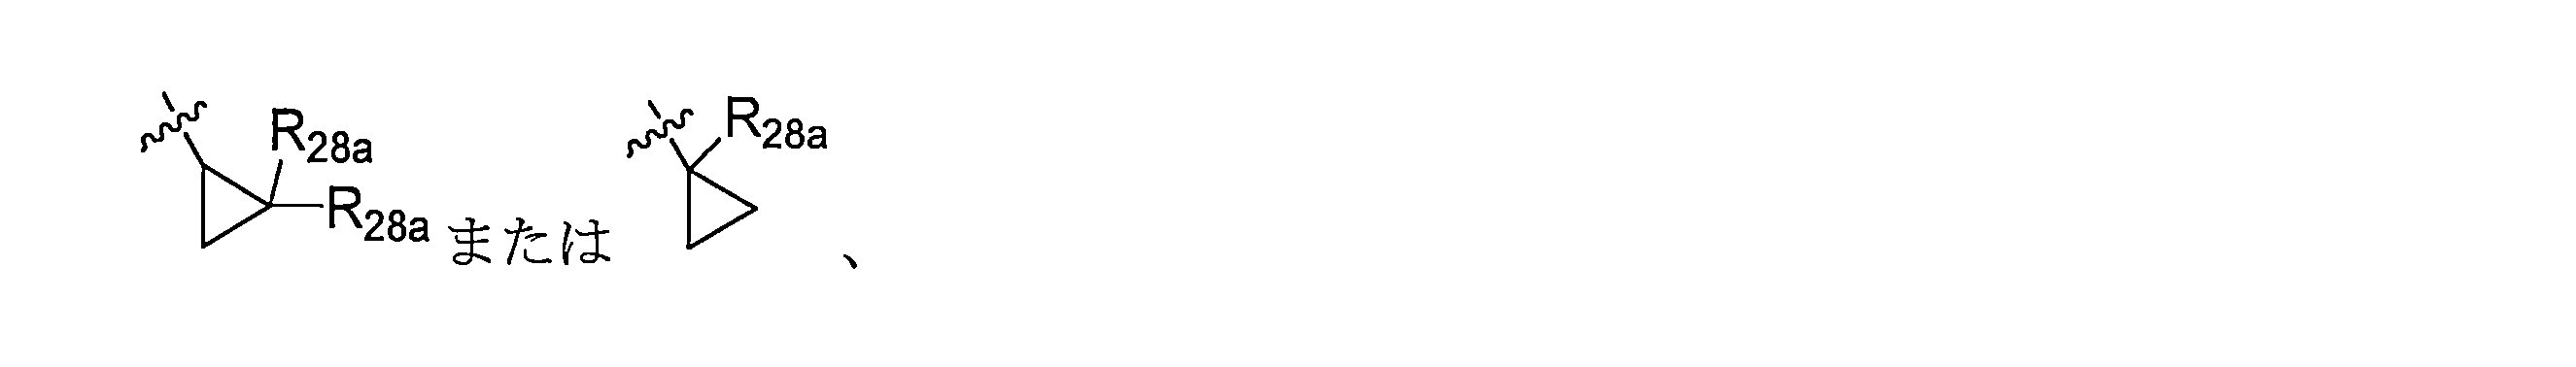
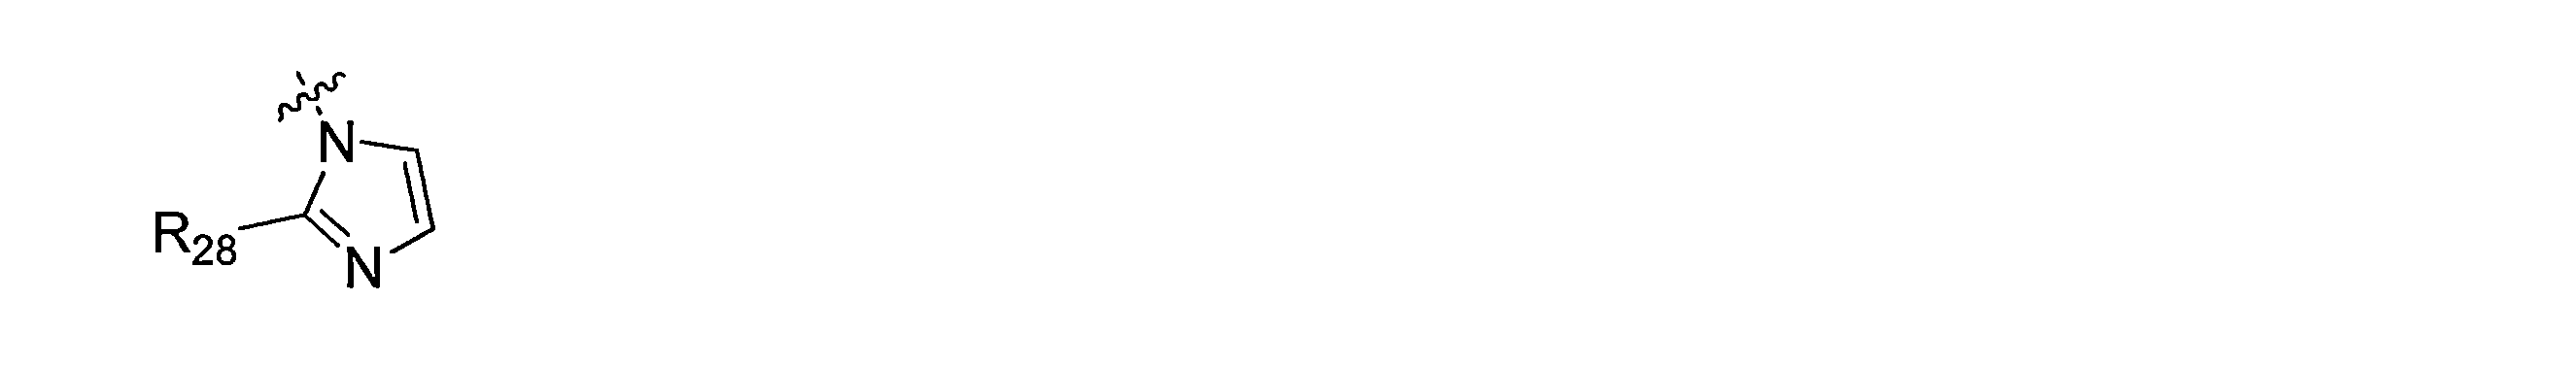
各R16は、独立して、ハロ、CN、任意に置換されたアルキル、任意に置換されたハロアルキル、ヒドロキシ、またはハロアルコキシから選択され、
各R26は、独立して、H、任意に置換されたアルキル、またはNR27aR27bであり、
各R27aおよびR27bは、独立して、H、任意に置換されたアルキル、任意に置換されたシクロアルキル(例えば、任意に置換された3~5員シクロアルキル)であるか、またはR27aおよびR27bは、それらが結合している窒素原子と一緒になって、4~6員ヘテロシクリルを形成し、
R28は、H、任意に置換されたアルキル、任意に置換されたアルコキシ、任意に置換されたヘテロアルキル、任意に置換されたアルキルアミン、任意に置換されたヒドロキシアルキル、アミン、任意に置換されたアルキニル、または任意に置換されたシクロアルキルであり、
oは、0、1、または2である。
In any of the aspects or embodiments described herein, the ULM has the following formula:
R1 is H, optionally substituted alkyl, or optionally substituted cycloalkyl;
R3 is an optionally substituted 5-6 membered heteroaryl;
W5 is optionally substituted phenyl, optionally substituted naphthyl, or optionally substituted pyridinyl;
one of R14a and R14b is H, optionally substituted alkyl, optionally substituted haloalkyl, optionally substituted alkoxy, optionally substituted hydroxylalkyl, optionally substituted alkylamine, optionally substituted heteroalkyl, optionally substituted alkyl-heterocycloalkyl, optionally substituted alkoxy-heterocycloalkyl, COR26, CONR27aR27b, NHCOR26, or NHCH3COR26; the other of R14a and R14b is H; or R14a, R14b together with the carbon atom to which they are attached form an optionally substituted 3- to 5-membered cycloalkyl, heterocycloalkyl, spirocycloalkyl, or spiroheterocyclyl, wherein the spiroheterocyclyl is not an epoxide or aziridine;
R15 is CN, optionally substituted fluoroalkyl,
each R16 is independently selected from halo, CN, optionally substituted alkyl, optionally substituted haloalkyl, hydroxy, or haloalkoxy;
each R26 is independently H, optionally substituted alkyl, or NR27aR27b;
each R27a and R27b is independently H, optionally substituted alkyl, optionally substituted cycloalkyl (e.g., optionally substituted 3- to 5-membered cycloalkyl), or R27a and R27b together with the nitrogen atom to which they are attached form a 4- to 6-membered heterocyclyl;
R28 is H, optionally substituted alkyl, optionally substituted alkoxy, optionally substituted heteroalkyl, optionally substituted alkylamine, optionally substituted hydroxyalkyl, amine, optionally substituted alkynyl, or optionally substituted cycloalkyl;
o is 0, 1, or 2.
本明細書に記載される態様または実施形態のいずれにおいても、ULMは、以下の式:

X4、X5、およびX6の各々が、CHおよびNから選択され、2個以下が、Nであり、
R1は、C1-6アルキルであり、
R14aおよびR14bのうちの一方が、H、任意に置換されたアルキル、任意に置換されたハロアルキル、任意に置換されたアルコキシ、任意に置換されたヒドロキシルアルキル、任意に置換されたアルキルアミン、任意に置換されたヘテロルキル、任意に置換されたアルキル-ヘテロシクロアルキル、任意に置換されたアルコキシ-ヘテロシクロアルキル、COR26、CONR27aR27b、NHCOR26、もしくはNHCH3COR26であり、R14aおよびR14bのうちの他方が、Hであり、またはR14aおよびR14bが、それらが結合している炭素原子と一緒になって、任意に置換された3~5員シクロアルキル、ヘテロシクロアルキル、スピロシクロアルキル、もしくはスピロヘテロシクリルを形成し、スピロヘテロシクリルが、エポキシドまたはアジリジンではなく、
各R27aおよびR27bは、独立して、HまたはC1-6アルキルもしくはシクロアルキル(例えば、3~5員シクロアルキル)であり、
qが、1、2、3、または4であり、
R15は、

R28は、H、メチル、CH2N(Me)2、CH2OH、CH2O(C1-4アルキル)、CH2NHC(O)C1-4アルキル、NH2、


R28Cは、H、メチル、フルオロ、またはクロロであり、
R16は、H、C1-4アルキル、フルオロ、クロロ、CN、またはC1-4アルコキシである。
In any of the aspects or embodiments described herein, the ULM has the following formula:
Each of X4, X5, and X6 is selected from CH and N, with up to 2 being N;
R1 is C1-6 alkyl;
one of R14a and R14b is H, optionally substituted alkyl, optionally substituted haloalkyl, optionally substituted alkoxy, optionally substituted hydroxylalkyl, optionally substituted alkylamine, optionally substituted heteroalkyl, optionally substituted alkyl-heterocycloalkyl, optionally substituted alkoxy-heterocycloalkyl, COR26, CONR27aR27b, NHCOR26, or NHCH3COR26; the other of R14a and R14b is H; or R14a and R14b together with the carbon atom to which they are attached form an optionally substituted 3- to 5-membered cycloalkyl, heterocycloalkyl, spirocycloalkyl, or spiroheterocyclyl, wherein the spiroheterocyclyl is not an epoxide or aziridine;
each R27a and R27b is independently H or C1-6 alkyl or cycloalkyl (e.g., 3- to 5-membered cycloalkyl);
q is 1, 2, 3, or 4;
R15 is,
R28 is H, methyl, CH2N(Me)2, CH2OH, CH2O(C1-4 alkyl), CH2NHC(O)C1-4 alkyl, NH2,
R28C is H, methyl, fluoro, or chloro;
R16 is H, C1-4 alkyl, fluoro, chloro, CN, or C1-4 alkoxy.
本明細書に記載される任意の態様または実施形態では、R14aおよびR14bは、H、C1-4アルキル、C1-4シクロアルキル、C1-4ハロアルキル、C1-4ヒドロキシアルキル、C1-4アルキルオキシアルキル、C1-4アルキル-NR27aR27b、およびCONR27aR27bから選択される。 In any aspect or embodiment described herein, R14a and R14b are selected from H, C1-4 alkyl, C1-4 cycloalkyl, C1-4 haloalkyl, C1-4 hydroxyalkyl, C1-4 alkyloxyalkyl, C1-4 alkyl-NR27aR27b, and CONR27aR27b.
本明細書に記載される任意の態様または実施形態では、R14aおよびR14bのうちの少なくとも一方は、Hである(例えば、R14aおよびR14bの両方は、Hである)。 In any aspect or embodiment described herein, at least one of R14a and R14b is H (e.g., both R14a and R14b are H).
本明細書に記載される任意の態様または実施形態では、R14aおよびR14bのうちの少なくとも一方は、任意に置換されたアルキル、任意に置換されたハロアルキル、任意に置換されたアルコキシ、任意に置換されたヒドロキシルアルキル、任意に置換されたアルキルアミン、任意に置換されたヘテロルキル、任意に置換されたアルキル-ヘテロシクロアルキル、任意に置換されたアルコキシ-ヘテロシクロアルキル、COR26、CONR27aR27b、NHCOR26、またはNHCH3COR26である。あるいは、本明細書に記載される任意の態様または実施形態では、R14aおよびR14bのうちの一方は、任意に置換されたアルキル、任意に置換されたハロアルキル、任意に置換されたアルコキシ、任意に置換されたヒドロキシルアルキル、任意に置換されたアルキルアミン、任意に置換されたヘテロルキル、任意に置換されたアルキルヘテロシクロアルキル、任意に置換されたアルコキシヘテロシクロアルキル、COR26、CONR27aR27b、NHCOR26、またはNHCH3COR26であり、R14aおよびR14bのうちの他方は、Hである。 In any aspect or embodiment described herein, at least one of R14a and R14b is an optionally substituted alkyl, an optionally substituted haloalkyl, an optionally substituted alkoxy, an optionally substituted hydroxylalkyl, an optionally substituted alkylamine, an optionally substituted heteroalkyl, an optionally substituted alkyl-heterocycloalkyl, an optionally substituted alkoxy-heterocycloalkyl, COR26, CONR27aR27b, NHCOR26, or NHCH3COR26. Alternatively, in any aspect or embodiment described herein, one of R14a and R14b is optionally substituted alkyl, optionally substituted haloalkyl, optionally substituted alkoxy, optionally substituted hydroxylalkyl, optionally substituted alkylamine, optionally substituted heteroalkyl, optionally substituted alkylheterocycloalkyl, optionally substituted alkoxyheterocycloalkyl, COR26, CONR27aR27b, NHCOR26, or NHCH3COR26, and the other of R14a and R14b is H.
本明細書に記載される任意の態様または実施形態では、R14aおよびR14bは、それらが結合している炭素原子と一緒になって、


本明細書に記載される態様または実施形態のいずれにおいても、R1は、C1-6アルキルである。 In any of the aspects or embodiments described herein, R1 is C1-6 alkyl.
本明細書に記載される態様または実施形態のいずれにおいても、R14aおよびR14bのうちの一方は、H、C1-6アルキル、C1-6ハロアルキル、任意に置換されたC1-4アルキルアミン、C1-6アルコキシ、(CH2)qC1-6アルコキシ、(CH2)qC1-6アルコキシ-C3-C7ヘテロシクロアルキル、(CH2)qOH、(CH2)qNR27aR27b、(CH2)qNHCOC1-6アルキル、C3-6シクロアルキル、またはNR27aR27bであり、各R26は、独立して、H、C1-6アルキル、またはNR27aR27bであり、各R27aおよびR27bは、独立して、HまたはC1-6アルキルであり、qは、1、2、3、または4である。 In any of the aspects or embodiments described herein, one of R14a and R14b is H, C1-6 alkyl, C1-6 haloalkyl, optionally substituted C1-4 alkylamine, C1-6 alkoxy, (CH2)qC1-6 alkoxy, (CH2)qC1-6 alkoxy-C3-C7 heterocycloalkyl, (CH2)qOH, (CH2)qNR27aR27b, (CH2)qNHCOC1-6 alkyl, C3-6 cycloalkyl, or NR27aR27b, each R26 is independently H, C1-6 alkyl, or NR27aR27b, each R27a and R27b is independently H or C1-6 alkyl, and q is 1, 2, 3, or 4.
本明細書に記載される態様または実施形態のいずれにおいても、R14aおよびR14bのうちの一方は、H、C1-4アルキル、C1-4ハロアルキル、C1-4アルコキシ、任意に置換されたC1-4アルキルアミン、(CH2)qC1-6アルコキシ、(CH2)qC1-6アルコキシ-C3-C7ヘテロシクロアルキル、(CH2)qOH、(CH2)qNR27aR27b、(CH2)qNHCOC1-6アルキル、C3-6シクロアルキル、またはNR27aR27bであり、各R26は、独立して、H、C1-4アルキル、またはNR27aR27bであり、各R27aおよびR27bは、独立して、HまたはC1-4アルキルであり、qは、1または2である。 In any of the aspects or embodiments described herein, one of R14a and R14b is H, C1-4 alkyl, C1-4 haloalkyl, C1-4 alkoxy, optionally substituted C1-4 alkylamine, (CH2)qC1-6 alkoxy, (CH2)qC1-6 alkoxy-C3-C7 heterocycloalkyl, (CH2)qOH, (CH2)qNR27aR27b, (CH2)qNHCOC1-6 alkyl, C3-6 cycloalkyl, or NR27aR27b, each R26 is independently H, C1-4 alkyl, or NR27aR27b, each R27a and R27b is independently H or C1-4 alkyl, and q is 1 or 2.
本明細書に記載される態様または実施形態のいずれにおいても、R28は、C1-6アルキル、C3-6シクロアルキル、C1-6ハロアルキル、(CH2)qOC1-6アルキル、(CH2)qOH、(CH2)qNR27aR27b、(CH2)qNHCOC1-6アルキル、または

R29は、H、C1-6アルキル、NR27aR27bまたはqNHCOC1-6アルキルであり、
式中、qは、1または2である。
In any of the aspects or embodiments described herein, R28 is C1-6 alkyl, C3-6 cycloalkyl, C1-6 haloalkyl, (CH2)qOC1-6 alkyl, (CH2)qOH, (CH2)qNR27aR27b, (CH2)qNHCOC1-6 alkyl, or
R29 is H, C1-6 alkyl, NR27aR27b or qNHCOC1-6 alkyl;
In the formula, q is 1 or 2.
本明細書に記載される態様または実施形態のいずれにおいても、R3は、イソキサゾリル、4-クロロイソキサゾリル、4-フルオロイソキサゾリル、またはピラゾリルである。本明細書に記載される態様または実施形態のいずれにおいても、Xは、CHである。 In any of the aspects or embodiments described herein, R3 is isoxazolyl, 4-chloroisoxazolyl, 4-fluoroisoxazolyl, or pyrazolyl. In any of the aspects or embodiments described herein, X is CH.
本明細書に記載される任意の態様または実施形態において、ULMは、以下の式、

式中、
R1、R14a、およびR14bは、本明細書に記載されるとおりであり、
Xは、CHまたはNであり、
R30は、H、F、またはClであり、
R16は、H、C1-4アルキル、フルオロ、クロロ、CN、またはC1-4アルコキシであり、
R28は、H、メチル、CH2N(Me)2、CH2OH、CH2O(C1-4アルキル)、CH2NHC(O)C1-4アルキル、NH2、

In the formula,
R1, R14a, and R14b are as described herein;
X is CH or N;
R30 is H, F, or Cl;
R16 is H, C1-4 alkyl, fluoro, chloro, CN, or C1-4 alkoxy;
R28 is H, methyl, CH2N(Me)2, CH2OH, CH2O(C1-4 alkyl), CH2NHC(O)C1-4 alkyl, NH2,
本明細書に記載される態様または実施形態のいずれにおいても、ULMは、以下の式、

本明細書に記載される態様または実施形態のいずれにおいても、ULMは、表1A、表1B、表1C、表1D、表2A、表2B、表2C、または表2Dに提供されるULMである。 In any aspect or embodiment described herein, the ULM is a ULM provided in Table 1A, Table 1B, Table 1C, Table 1D, Table 2A, Table 2B, Table 2C, or Table 2D.
本明細書に記載される態様または実施形態のいずれにおいても、化合物は、本明細書に記載されるリンカー(L)を含む。 In any of the aspects or embodiments described herein, the compound includes a linker (L) as described herein.
本明細書に記載される態様または実施形態のいずれにおいても、リンカー(L)は、1~10個のエチレングリコール単位を含むアリールまたはフェニルで任意に置換されたポリエチレンオキシ基である。 In any of the aspects or embodiments described herein, the linker (L) is a polyethyleneoxy group optionally substituted with aryl or phenyl containing 1 to 10 ethylene glycol units.
本明細書に記載される態様または実施形態のいずれにおいても、化合物は、表1A、表1B、表1C、表1D、表2A、表2B、表2C、および表2Dの化合物から選択される(例えば、化合物1~765から選択される)。 In any of the aspects or embodiments described herein, the compound is selected from the compounds of Tables 1A, 1B, 1C, 1D, 2A, 2B, 2C, and 2D (e.g., selected from compounds 1-765).
本明細書に記載される態様または実施形態のいずれにおいても、化合物は、80%以上のD最大を有する。 In any of the aspects or embodiments described herein, the compound has a Dmax of 80% or greater.
追加の態様では、説明は、有効量の本明細書に記載される二官能性化合物と、薬学的に許容される担体とを含む組成物を提供する。 In an additional aspect, the description provides a composition comprising an effective amount of a bifunctional compound described herein and a pharma- ceutically acceptable carrier.
本明細書に記載される態様または実施形態のいずれにおいても、本明細書に記載される組成物は、追加の生体活性剤または請求項1~32のいずれかに記載の別の二官能性化合物のうちの少なくとも1つをさらに含む。 In any of the aspects or embodiments described herein, the compositions described herein further comprise at least one additional bioactive agent or another bifunctional compound described in any of claims 1-32.
本明細書に記載される態様または実施形態のいずれにおいても、追加の生体活性剤は、抗がん剤である。 In any of the aspects or embodiments described herein, the additional bioactive agent is an anticancer agent.
本明細書に記載の態様または実施形態のいずれにおいても、本明細書に記載される組成物は、薬学的に許容される担体と、対象における疾患または障害を治療するための有効量の本明細書に記載される少なくとも1つの化合物とを含み、方法は、組成物を、それを必要とする対象に投与することを含み、化合物は、疾患または障害の少なくとも1つの症状を治療または改善するのに有効である。 In any of the aspects or embodiments described herein, the compositions described herein include a pharma- ceutically acceptable carrier and an effective amount of at least one compound described herein to treat a disease or disorder in a subject, and the methods include administering the composition to a subject in need thereof, wherein the compound is effective to treat or ameliorate at least one symptom of the disease or disorder.
本明細書に記載される任意の態様または実施形態において、疾患または障害は、SMARCA1、BRAHMA、またはBRMの蓄積および凝集に関連する。本明細書に記載される任意の態様または実施形態において、疾患または障害は、がんである。本明細書に記載される任意の態様または実施形態では、がんは、SWI/SNF関連がん、またはSMARCA4変異を有するがん(例えば、肺がんまたは非小細胞肺がん)である。本明細書に記載される任意の態様または実施形態では、がんは、SMARCA4欠損がん、または正常なSMARCA4発現と比較して(例えば、野生型SMARCA4を有する同様の位置にある非がん細胞における非変異SMARCA4またはSMARCA4の発現と比較して)SMARCA4の発現が減少したがん、例えば、肺がんまたは非小細胞肺がんである。 In any aspect or embodiment described herein, the disease or disorder is associated with accumulation and aggregation of SMARCA1, BRAHMA, or BRM. In any aspect or embodiment described herein, the disease or disorder is cancer. In any aspect or embodiment described herein, the cancer is a SWI/SNF-associated cancer, or a cancer with a SMARCA4 mutation (e.g., lung cancer or non-small cell lung cancer). In any aspect or embodiment described herein, the cancer is a SMARCA4-deficient cancer, or a cancer with reduced expression of SMARCA4 compared to normal SMARCA4 expression (e.g., compared to non-mutated SMARCA4 or SMARCA4 expression in similarly located non-cancer cells with wild-type SMARCA4), e.g., lung cancer or non-small cell lung cancer.
アッセイおよび分解データ
SW1573細胞におけるBRM分解のウェスタンブロットスクリーニング
BRM分解(D最大およびDC50)を評価するために、細胞を、ウェル当たり180μLのDMEM増殖培地(1%のpen-strep、1%のHEPES、および10%のFBSを含む)中の96ウェルブラック/クリア底プレート中、8000個/ウェルで播種した。プレートを一晩インキュベートして接着を可能にした。翌朝、20μLの10倍標的化合物濃度(1%DMSO)を、適切なウェルに添加することによって細胞を処理し、インキュベーターに一晩(18~20時間)戻した。最終DMSO濃度は0.1%であった。
Assay and Degradation Data Western Blot Screening of BRM Degradation in SW1573 Cells To assess BRM degradation (Dmax and DC50), cells were seeded at 8000 cells/well in 96-well black/clear bottom plates in 180 μL of DMEM growth medium (containing 1% pen-strep, 1% HEPES, and 10% FBS) per well. Plates were incubated overnight to allow attachment. The following morning cells were treated by adding 20 μL of 10x target compound concentration (1% DMSO) to the appropriate wells and returned to the incubator overnight (18-20 hours). Final DMSO concentration was 0.1%.
溶解のために、付着細胞を100μLのDPBSで1回洗浄した。細胞を、氷上で40μLの1X RIPA+HALTプロテアーゼ阻害剤中で10分間溶解し、-80℃で使用するまで凍結した。解凍された溶解物を、1.2μmのフィルタープレートでの濾過によって洗浄するか、または代替的に、2300gで4℃において30分間回転洗浄した。 For lysis, adherent cells were washed once with 100 μL DPBS. Cells were lysed in 40 μL 1X RIPA + HALT protease inhibitors on ice for 10 min and frozen at -80°C until use. Thawed lysates were washed by filtration through a 1.2 μm filter plate or alternatively spun at 2300 g for 30 min at 4°C.
ブロットするために、各ウェスタン試料について、30μLの溶解物を10μLの4倍LDS試料緩衝液に添加し、次いで、サーマルサイクラー中で、95℃で5分間変性させ、氷上に配置した。試料を、4~15%のトリス/グリシンゲルに充填し、各ブロットについて充填した4μLのラダーおよび12μLの各試料とともに1倍トリス/グリシン緩衝液中で、250定常ボルトで25分間実行した。タンパク質を、Turbo/midiデフォルトプログラムを用いたBioRad Turboドライトランスファーユニットを用いて、ゲルからNCに移した。すべてのブロットをddH2Oですすぎ、ロッカー上のTBST(0.1%)中5%BSA中、室温で1時間ブロックした。ブロットを、ロッカー(BRMに対して1:1000(Cell Signaling Tech.カタログ番号11966)およびアルファ-チューブリンに対して1:2000(Cell Signaling Tech.カタログ番号3873)、対照タンパク質上で、4℃で一晩、TBST(0.1%)中の5%BSA中の一次抗体に曝露した。ブロットを、室温において、ロッカー上でTBST(0.1%)で5分間、3回洗浄した。二次抗体を加え、ブロットを、TBST(0.1%)中の5%BSA中の1:18,000抗ウサギ-HRPおよび/または抗マウス-HRPと、室温ロッカー上で1時間インキュベートした。ブロットを、ロッカー上で室温において、TBST(0.1%)中で5分間、3回洗浄した。信号は、Femto Maximum Sensitivity基板を用いて4分間現像し、ChemiDoc(商標)でブロットを読み取った。 To blot, for each western sample, 30 μL of lysate was added to 10 μL of 4x LDS sample buffer, then denatured in a thermal cycler at 95°C for 5 minutes and placed on ice. Samples were loaded onto a 4-15% Tris/Glycine gel and run at 250 constant volts for 25 minutes in 1x Tris/Glycine buffer with 4 μL of ladder and 12 μL of each sample loaded for each blot. Proteins were transferred from the gel to NC using a BioRad Turbo dry transfer unit using the Turbo/midi default program. All blots were rinsed with ddH2O and blocked in 5% BSA in TBST (0.1%) on a rocker for 1 hour at room temperature. Blots were exposed to primary antibodies in 5% BSA in TBST (0.1%) on a rocker (1:1000 for BRM (Cell Signaling Tech. Cat. No. 11966) and 1:2000 for alpha-tubulin (Cell Signaling Tech. Cat. No. 3873), control proteins, overnight at 4°C. Blots were washed 3 times for 5 min with TBST (0.1%) on a rocker at room temperature. Secondary antibodies were added and blots were incubated with 1:18,000 anti-rabbit-HRP and/or anti-mouse-HRP in 5% BSA in TBST (0.1%) for 1 hr on a rocker at room temperature. Blots were washed 3 times for 5 min in TBST (0.1%) on a rocker at room temperature. Signals were analyzed using a Femto Maximum The blot was developed with Sensitivity substrate for 4 minutes and read on a ChemiDoc™.
SW1573細胞におけるBRM分解の細胞内ウエスタンスクリーニング
BRM分解(D最大およびDC50)を評価するために、細胞を、ウェル当たり180μLのDMEM増殖培地(1%のpen-strep、1%のHEPES、および10%のFBSを含む)中の96ウェルブラック/クリア底プレート中、8000個/ウェルで播種した。プレートを一晩インキュベートして接着を可能にした。翌朝、20μLの10倍化合物(1%DMSO)を、適切なウェルに添加することによって細胞を処理し、インキュベーターに一晩(18~20時間)戻した。最終DMSO濃度は0.1%であった。
Intracellular Western Screening of BRM Degradation in SW1573 Cells To assess BRM degradation (Dmax and DC50), cells were seeded at 8000 cells/well in 96-well black/clear bottom plates in 180 μL of DMEM growth medium (containing 1% pen-strep, 1% HEPES, and 10% FBS) per well. Plates were incubated overnight to allow attachment. The following morning cells were treated by adding 20 μL of 10x compound (1% DMSO) to the appropriate wells and returned to the incubator overnight (18-20 hours). Final DMSO concentration was 0.1%.
処理のために、プレートをインキュベーターから取り出し、培地を除去し、直ちに200μLの冷(4℃)DPBSをすべてのウェルに添加した。次いで、DPBSを除去し、DPBS(4℃)中の50μLの4%パラホルムアルデヒド(PFA)をすべてのウェルに添加し、プレートを室温で20分間インキュベートした。次いで、PFAを除去し、0.5%Triton X-100を含む200μLのTBS-Tをすべてのウェルに添加し、プレートを室温で30分間インキュベートした。次いで、0.5%Triton X-100を含むTBS-Tを除去し、50μLのLi-Cor遮断溶液を添加し、プレートを室温で少なくとも1時間インキュベートした。遮断溶液を除去し、対照タンパク質である一次抗体カクテルを含む50μLのLi-Cor遮断溶液を添加した(BRM(Cell Signaling Tech.カタログ番号11966)については1:1000、およびアルファ-チューブリン(Sigmaカタログ番号T6074)については1:2000) 次いで、プレートを翌日まで冷たい部屋に配置した。 For treatment, plates were removed from the incubator, medium was removed, and 200 μL of cold (4°C) DPBS was immediately added to all wells. DPBS was then removed, 50 μL of 4% paraformaldehyde (PFA) in DPBS (4°C) was added to all wells, and plates were incubated at room temperature for 20 minutes. PFA was then removed, 200 μL of TBS-T with 0.5% Triton X-100 was added to all wells, and plates were incubated at room temperature for 30 minutes. TBS-T with 0.5% Triton X-100 was then removed, 50 μL of Li-Cor blocking solution was added, and plates were incubated at room temperature for at least 1 hour. The blocking solution was removed and 50 μL of Li-Cor blocking solution was added containing a primary antibody cocktail with control proteins (1:1000 for BRM (Cell Signaling Tech. Cat. No. 11966) and 1:2000 for alpha-tubulin (Sigma Cat. No. T6074)). The plate was then placed in a cold room until the next day.
翌日、プレートをTBS-T、ウェル当たり200μLで3回洗浄した。LI-COR遮断溶液(抗ウサギ_800nmおよび抗マウス_680nm)中50μLの二次抗体カクテルを、すべてのウェルに添加した(1:5000で希釈)。プレートを、光から保護しながら、室温で少なくとも1時間インキュベートした。プレートをTBS-Tで2回、ウェル当たり200μLで洗浄した。 The next day, plates were washed 3 times with TBS-T, 200 μL per well. 50 μL of secondary antibody cocktail in LI-COR blocking solution (anti-rabbit_800 nm and anti-mouse_680 nm) was added to all wells (diluted 1:5000). Plates were incubated at room temperature for at least 1 hour, protected from light. Plates were washed 2 times with TBS-T, 200 μL per well.
各プレートを読み取るために、TBS-Tを取り外し、各プレートを反転させてペーパータオル上にタップした。プレートをLI-COR Odyssey上で読み取り、両方のチャネルのデフォルト強度設定は5.0であった。LI-COR画像を、Image Studio Liteの細胞内ウェスタン特徴を用いて分析した。 To read each plate, the TBS-T was removed and each plate was inverted and tapped onto a paper towel. Plates were read on a LI-COR Odyssey with default intensity settings of 5.0 for both channels. LI-COR images were analyzed using the intracellular Western feature in Image Studio Lite.
以下の化合物は、上に記載される条件下で試験した場合、標的タンパク質分解を示した。

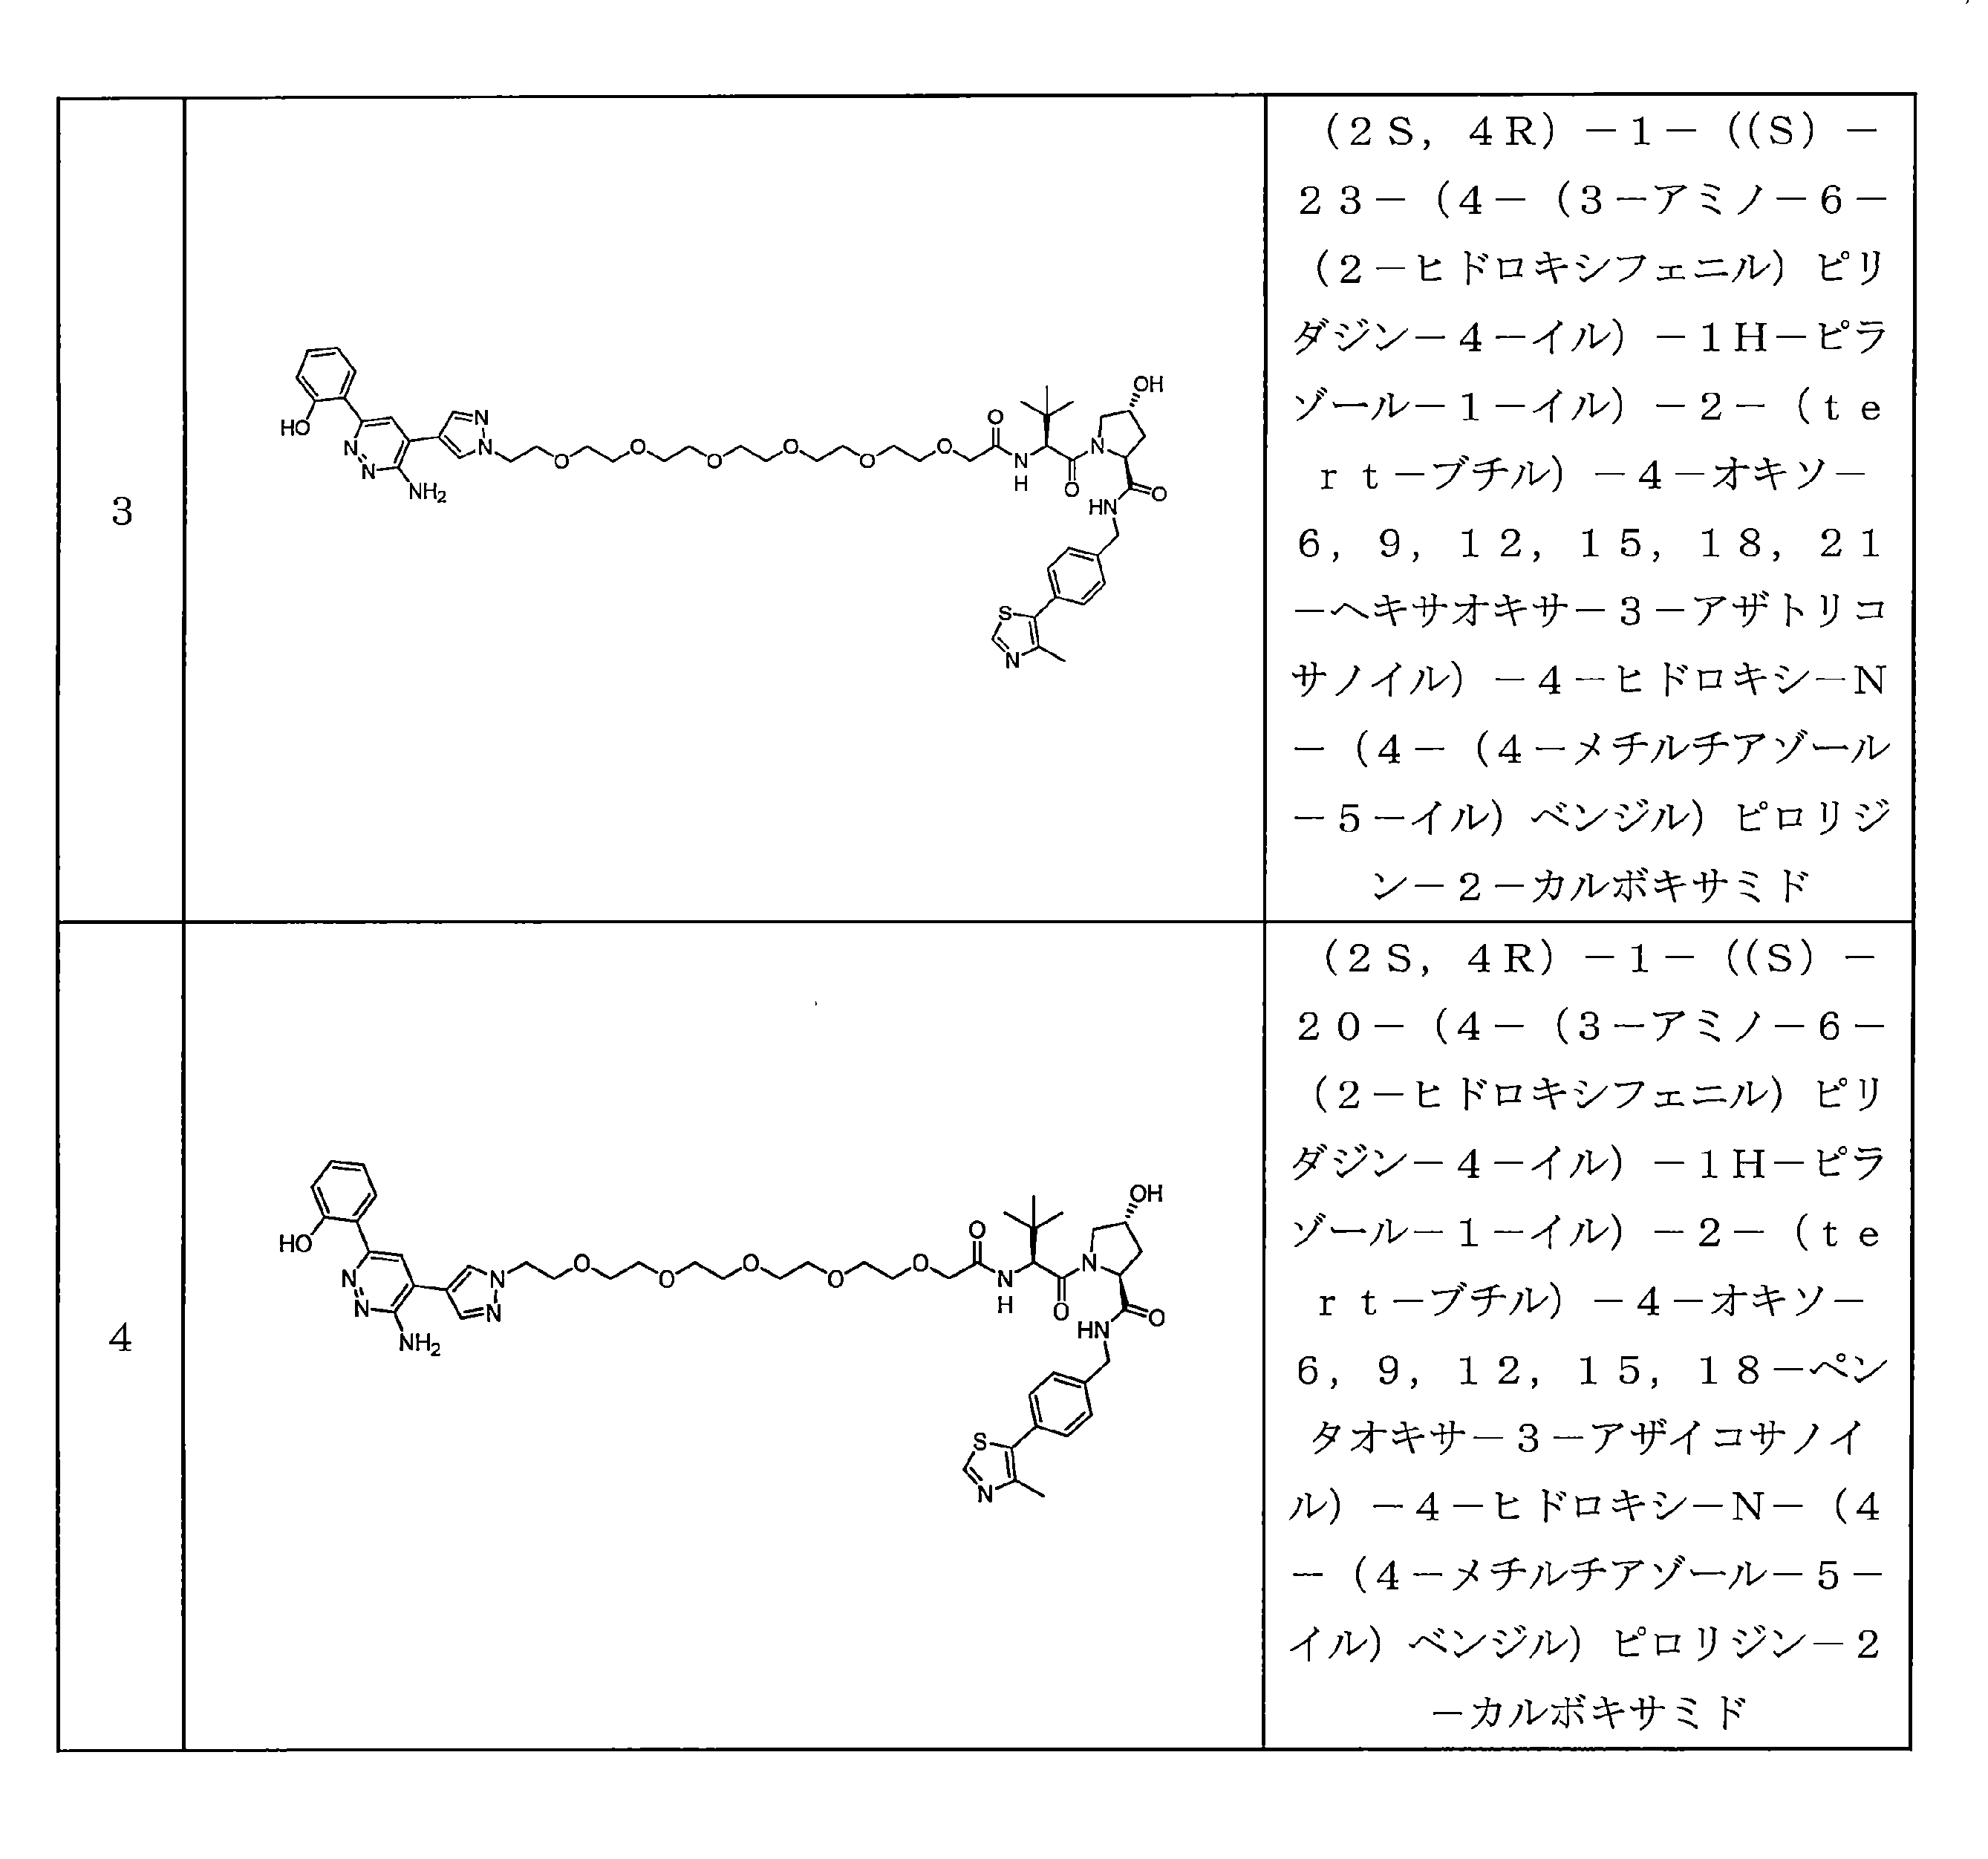


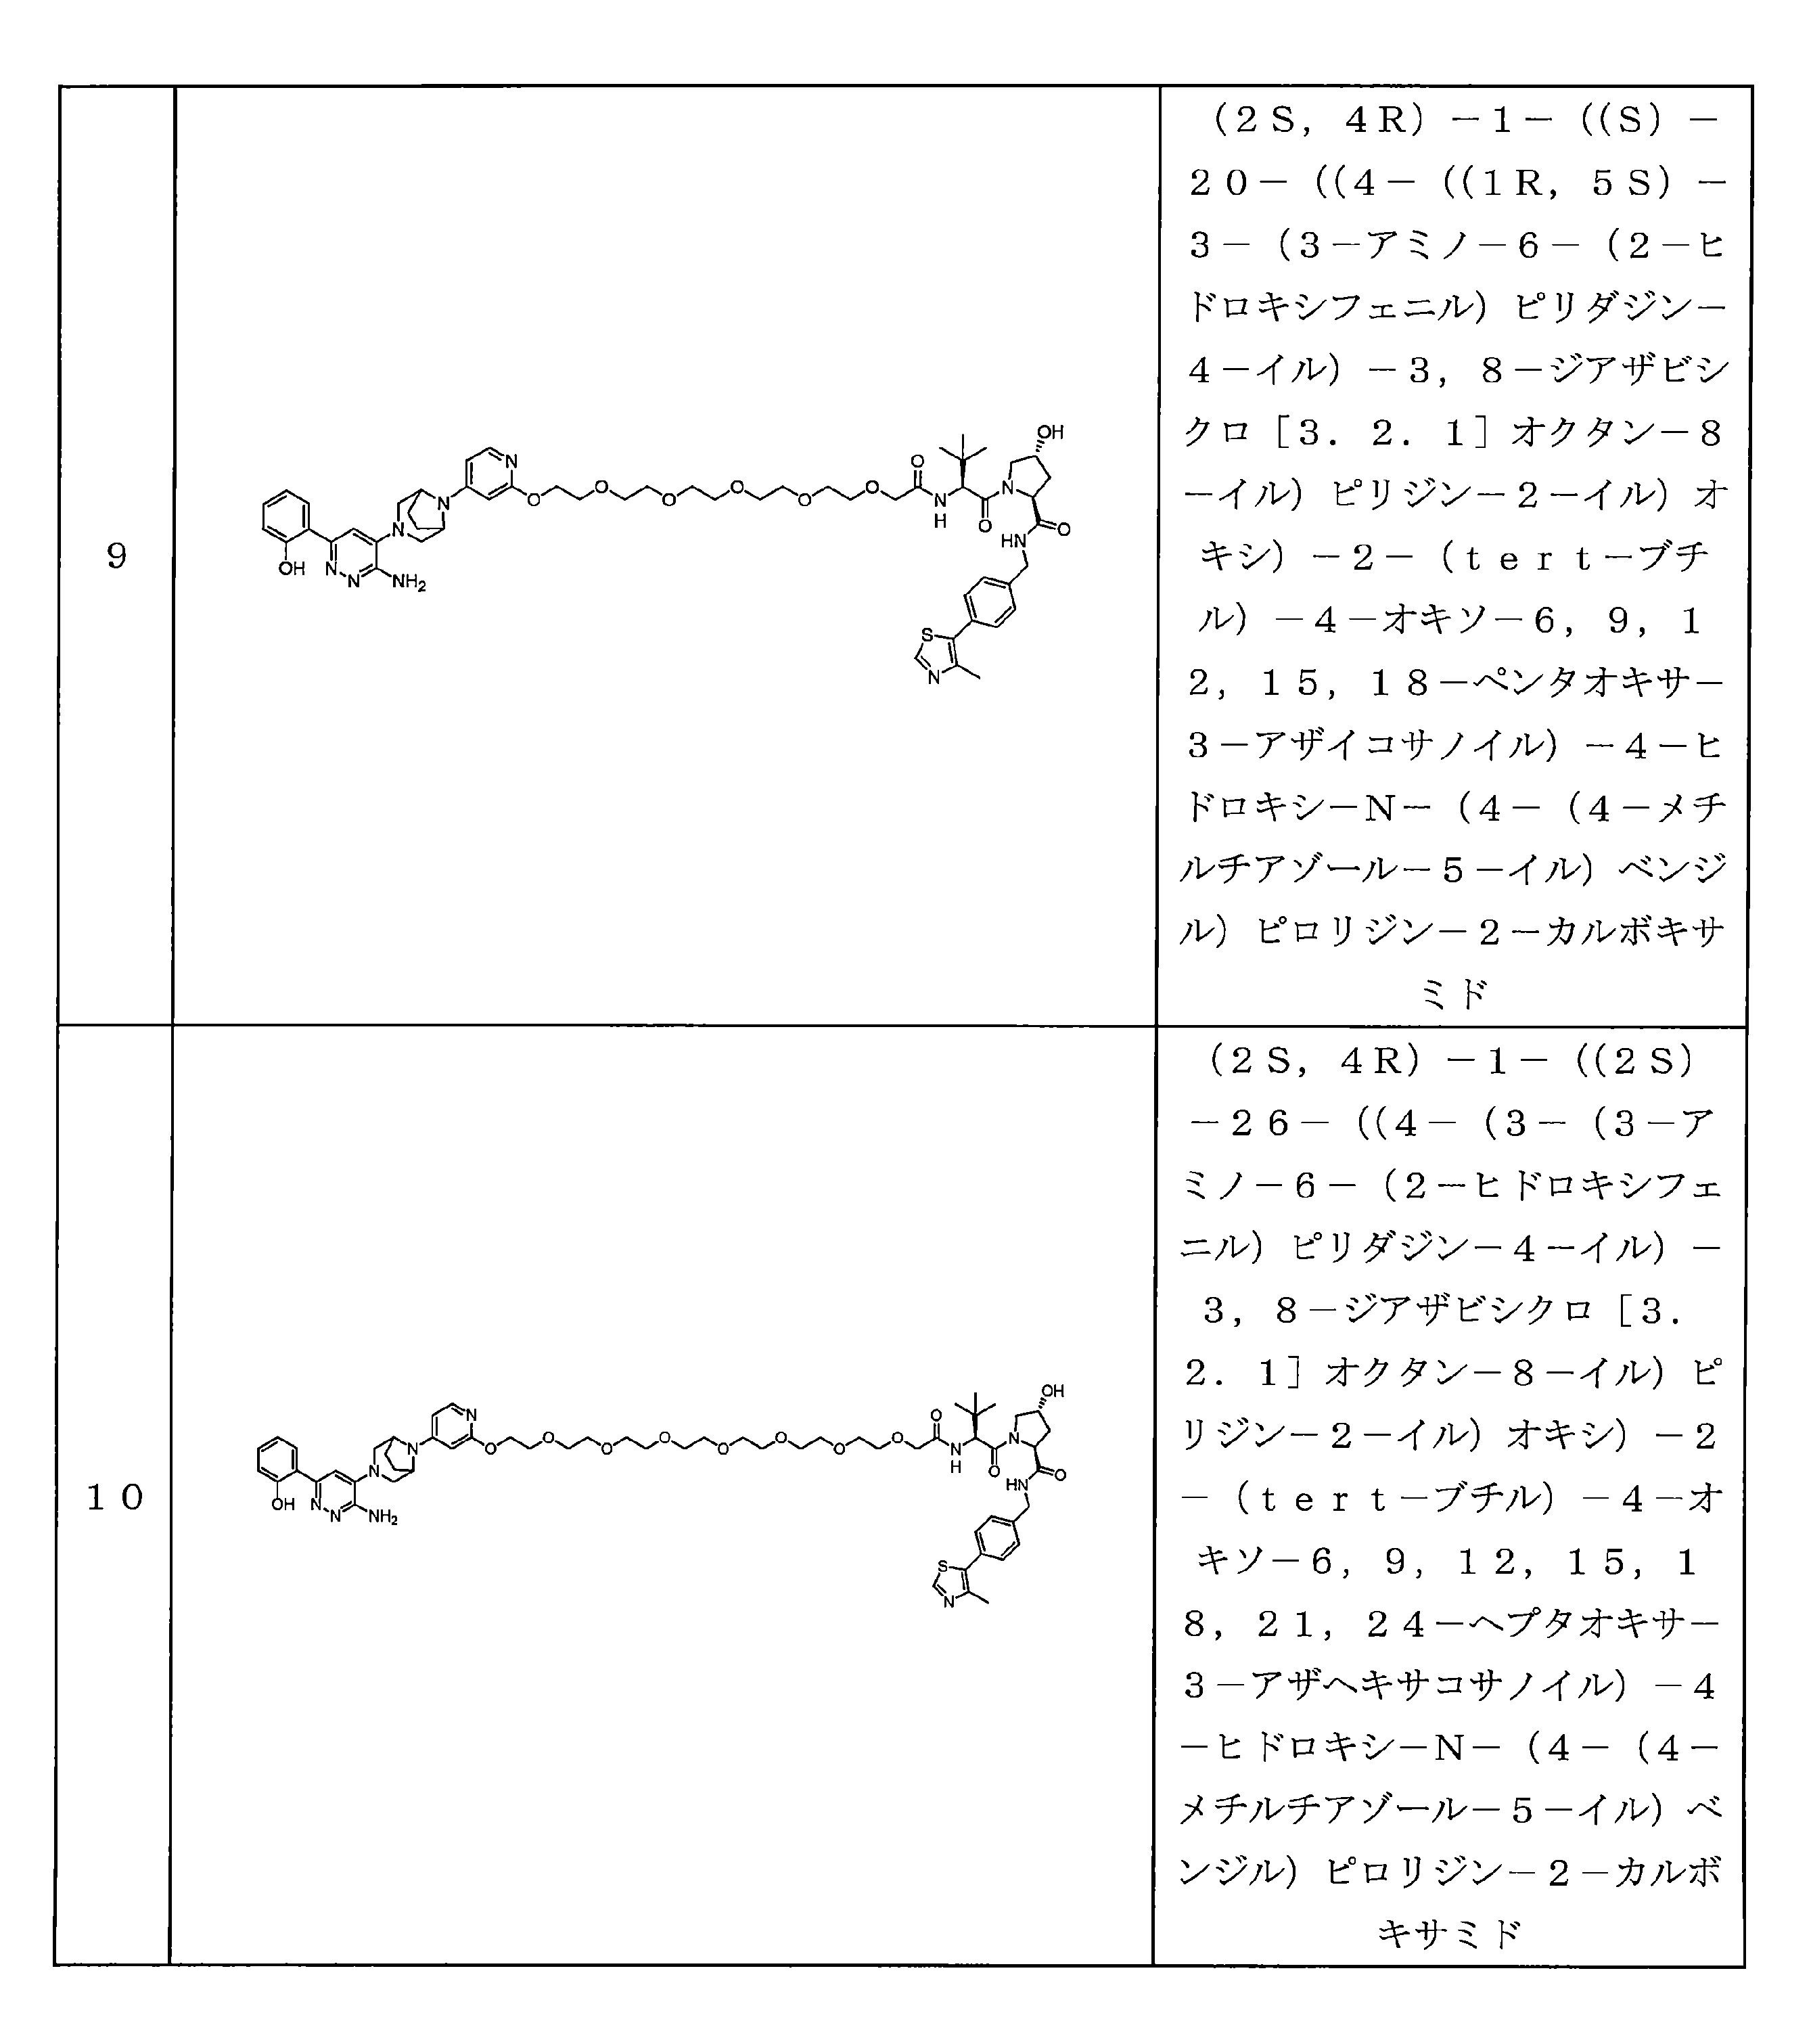
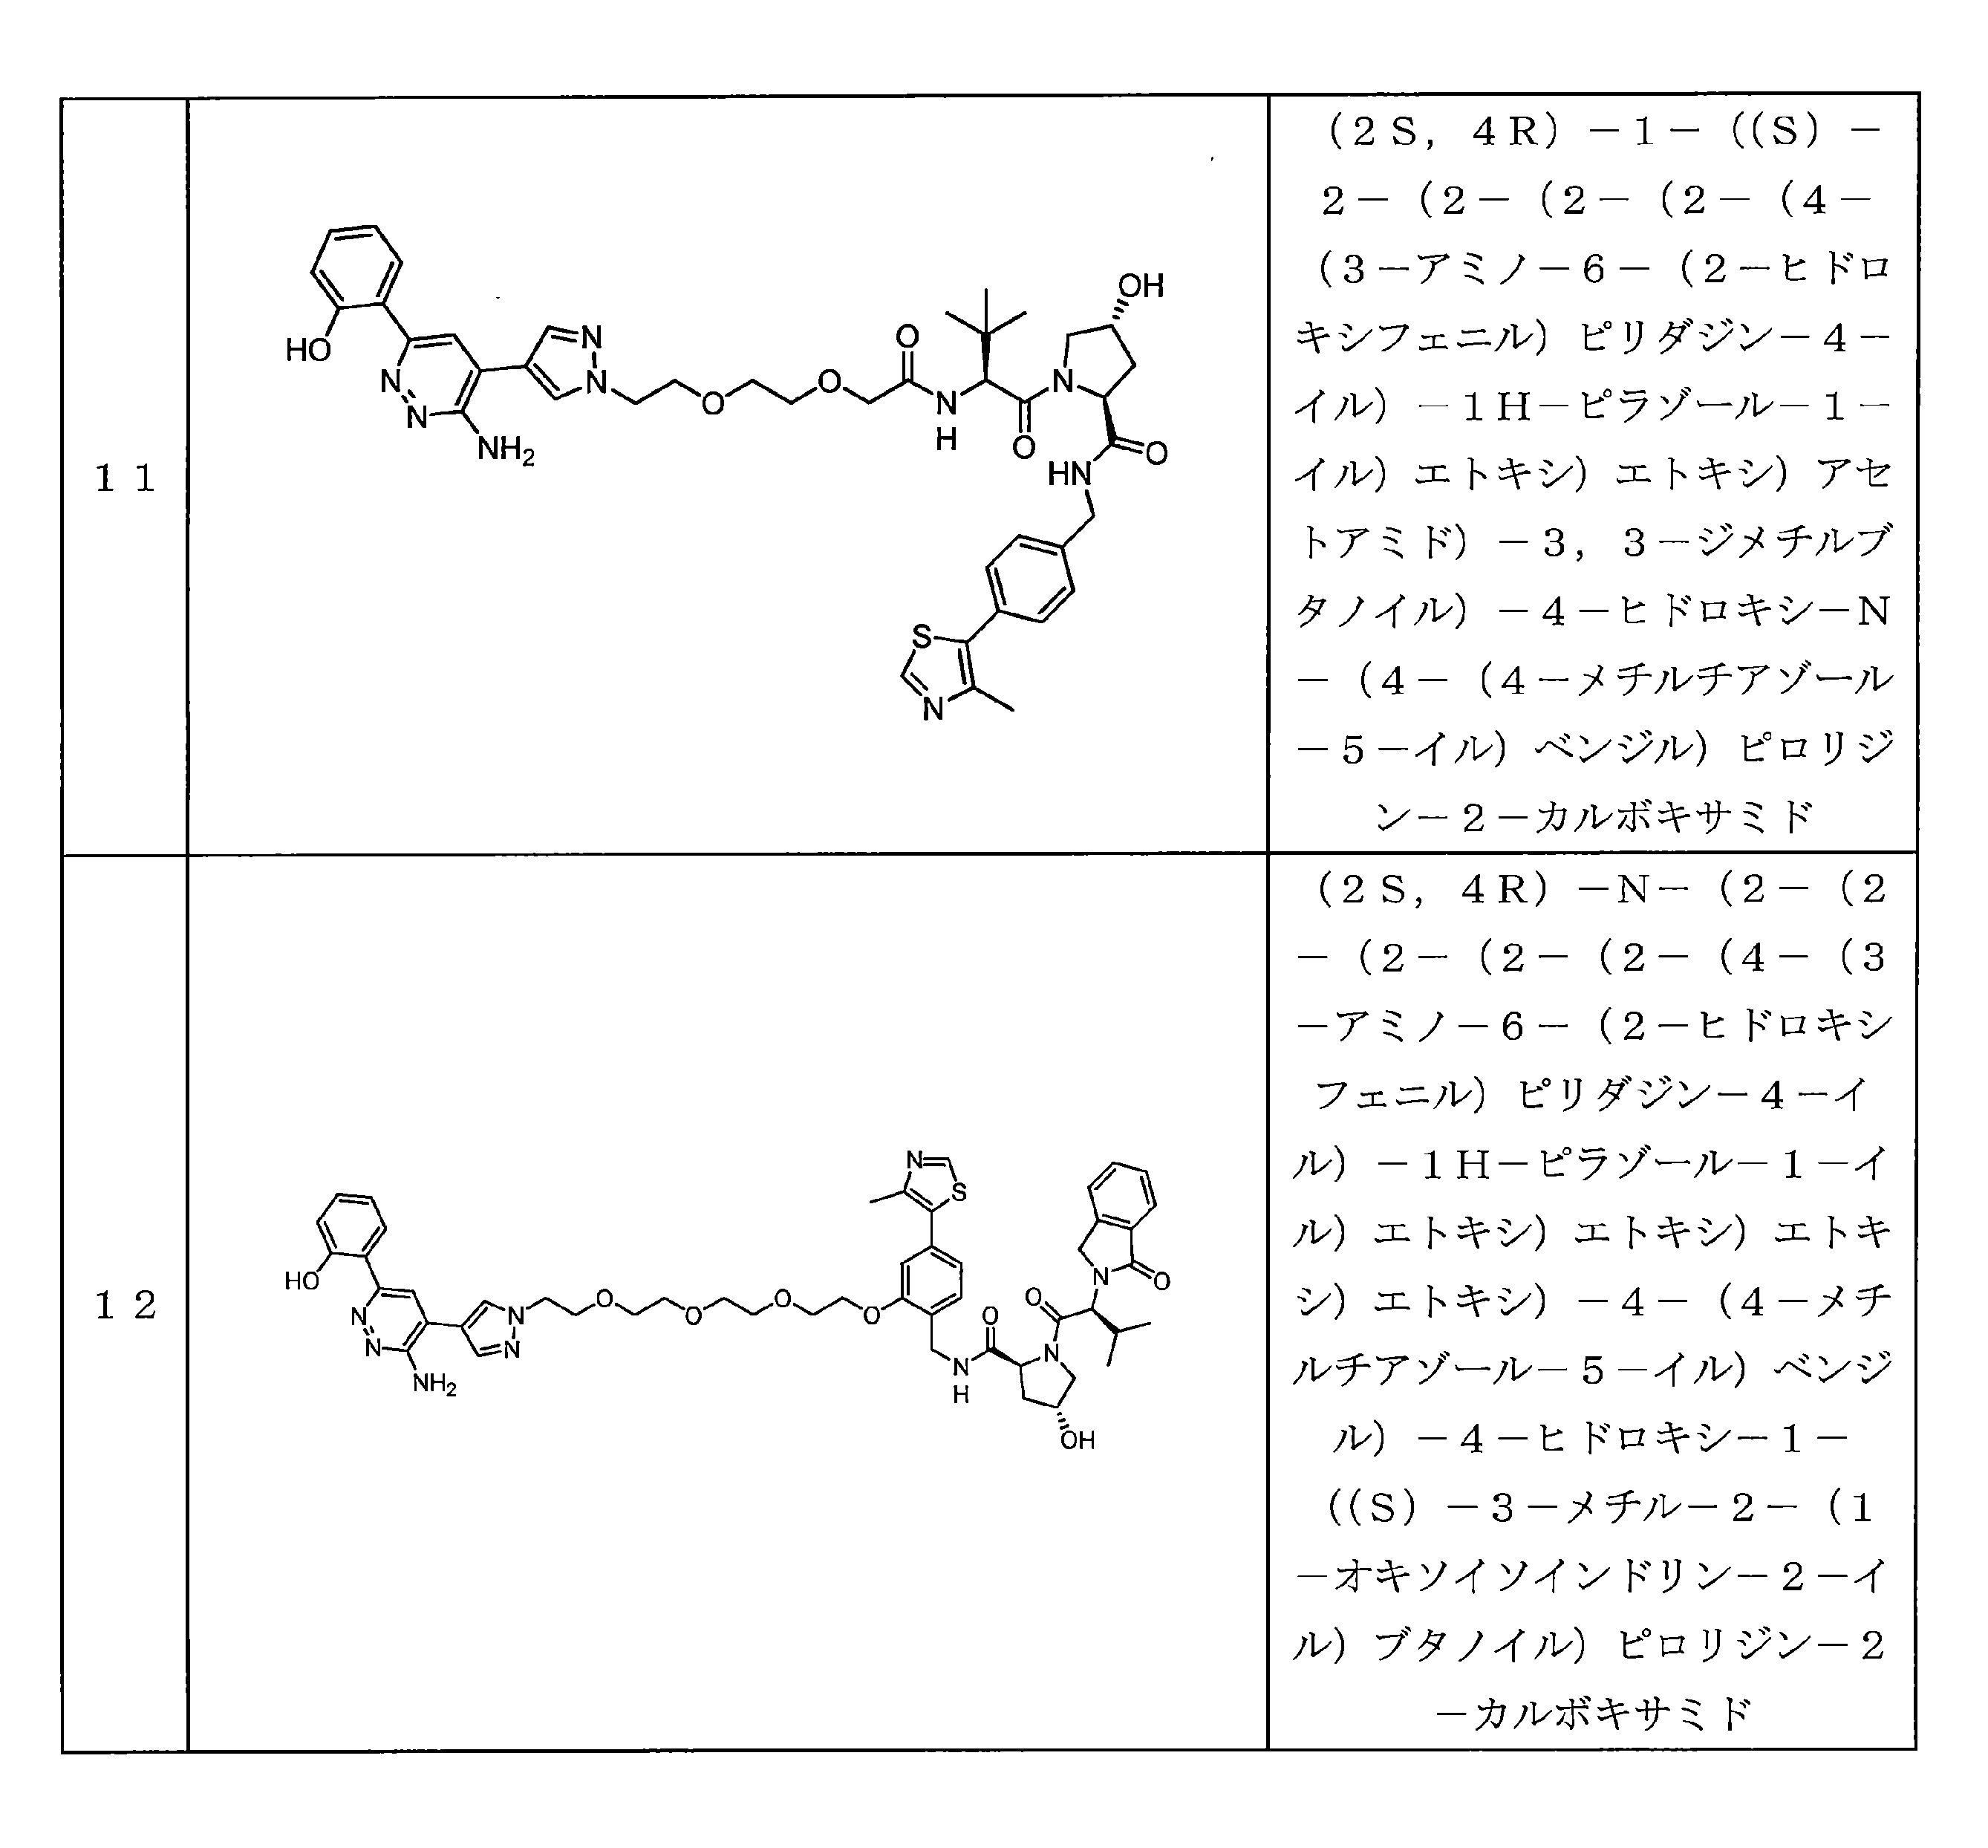

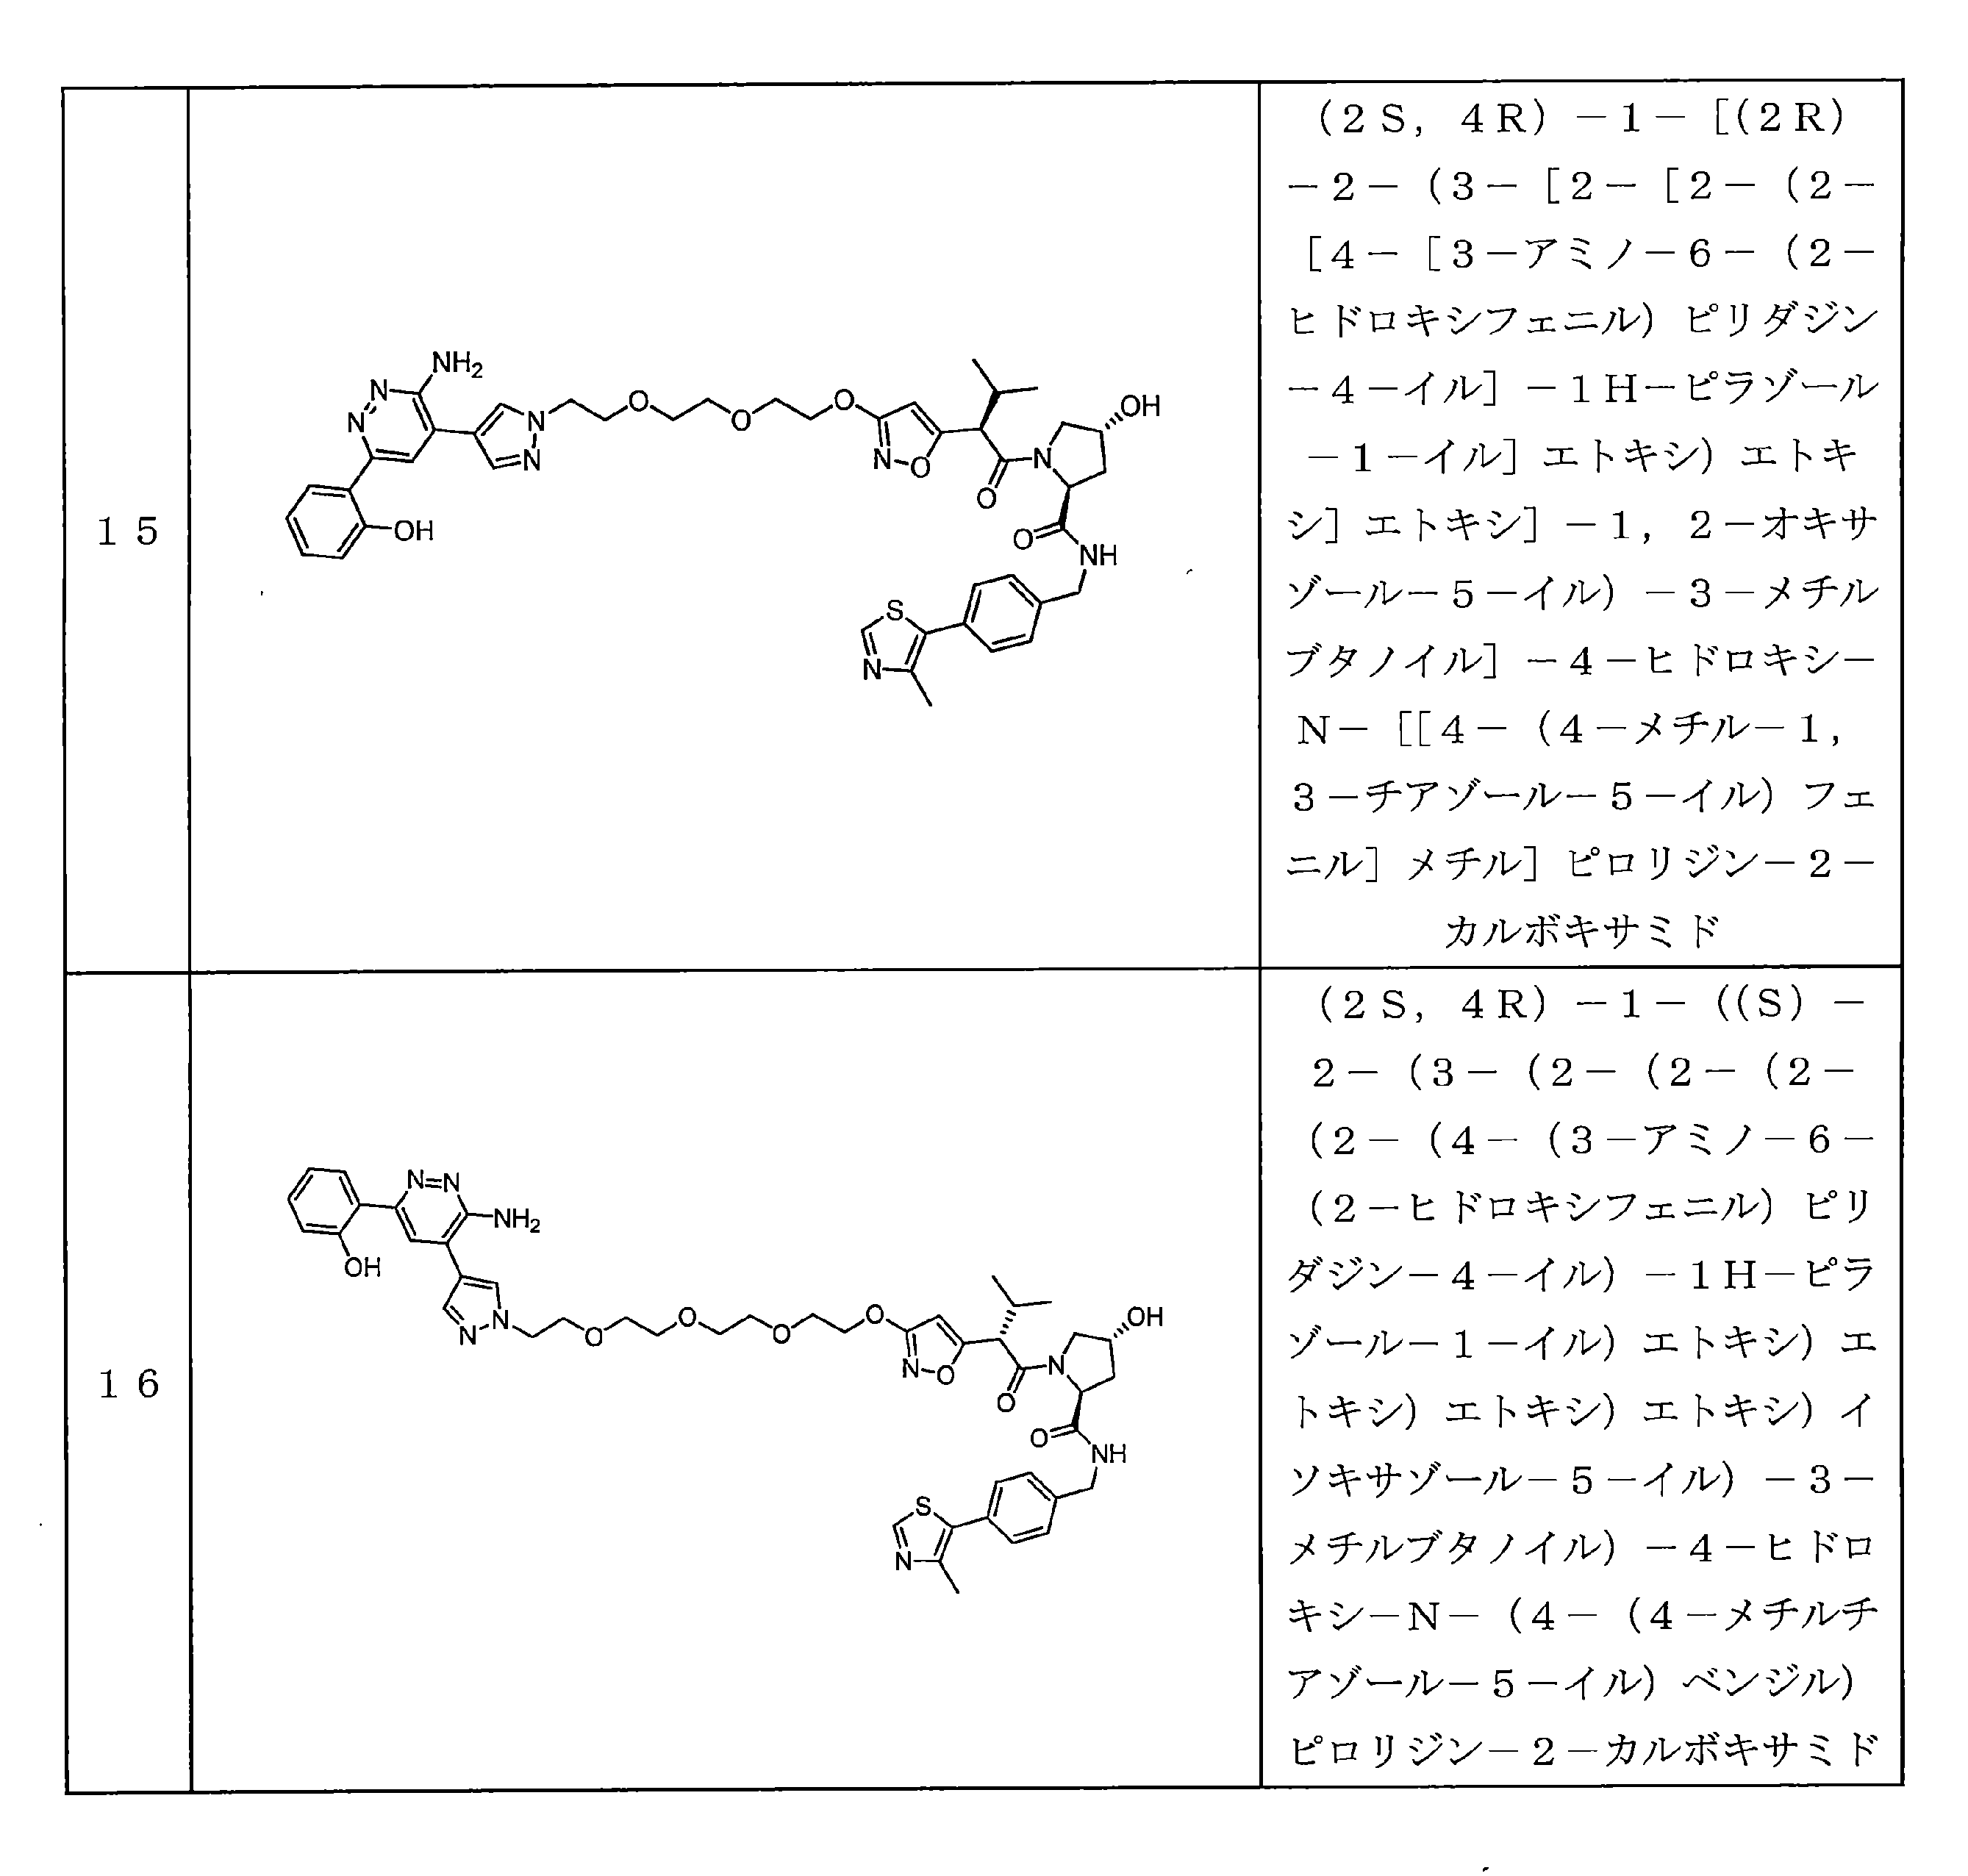
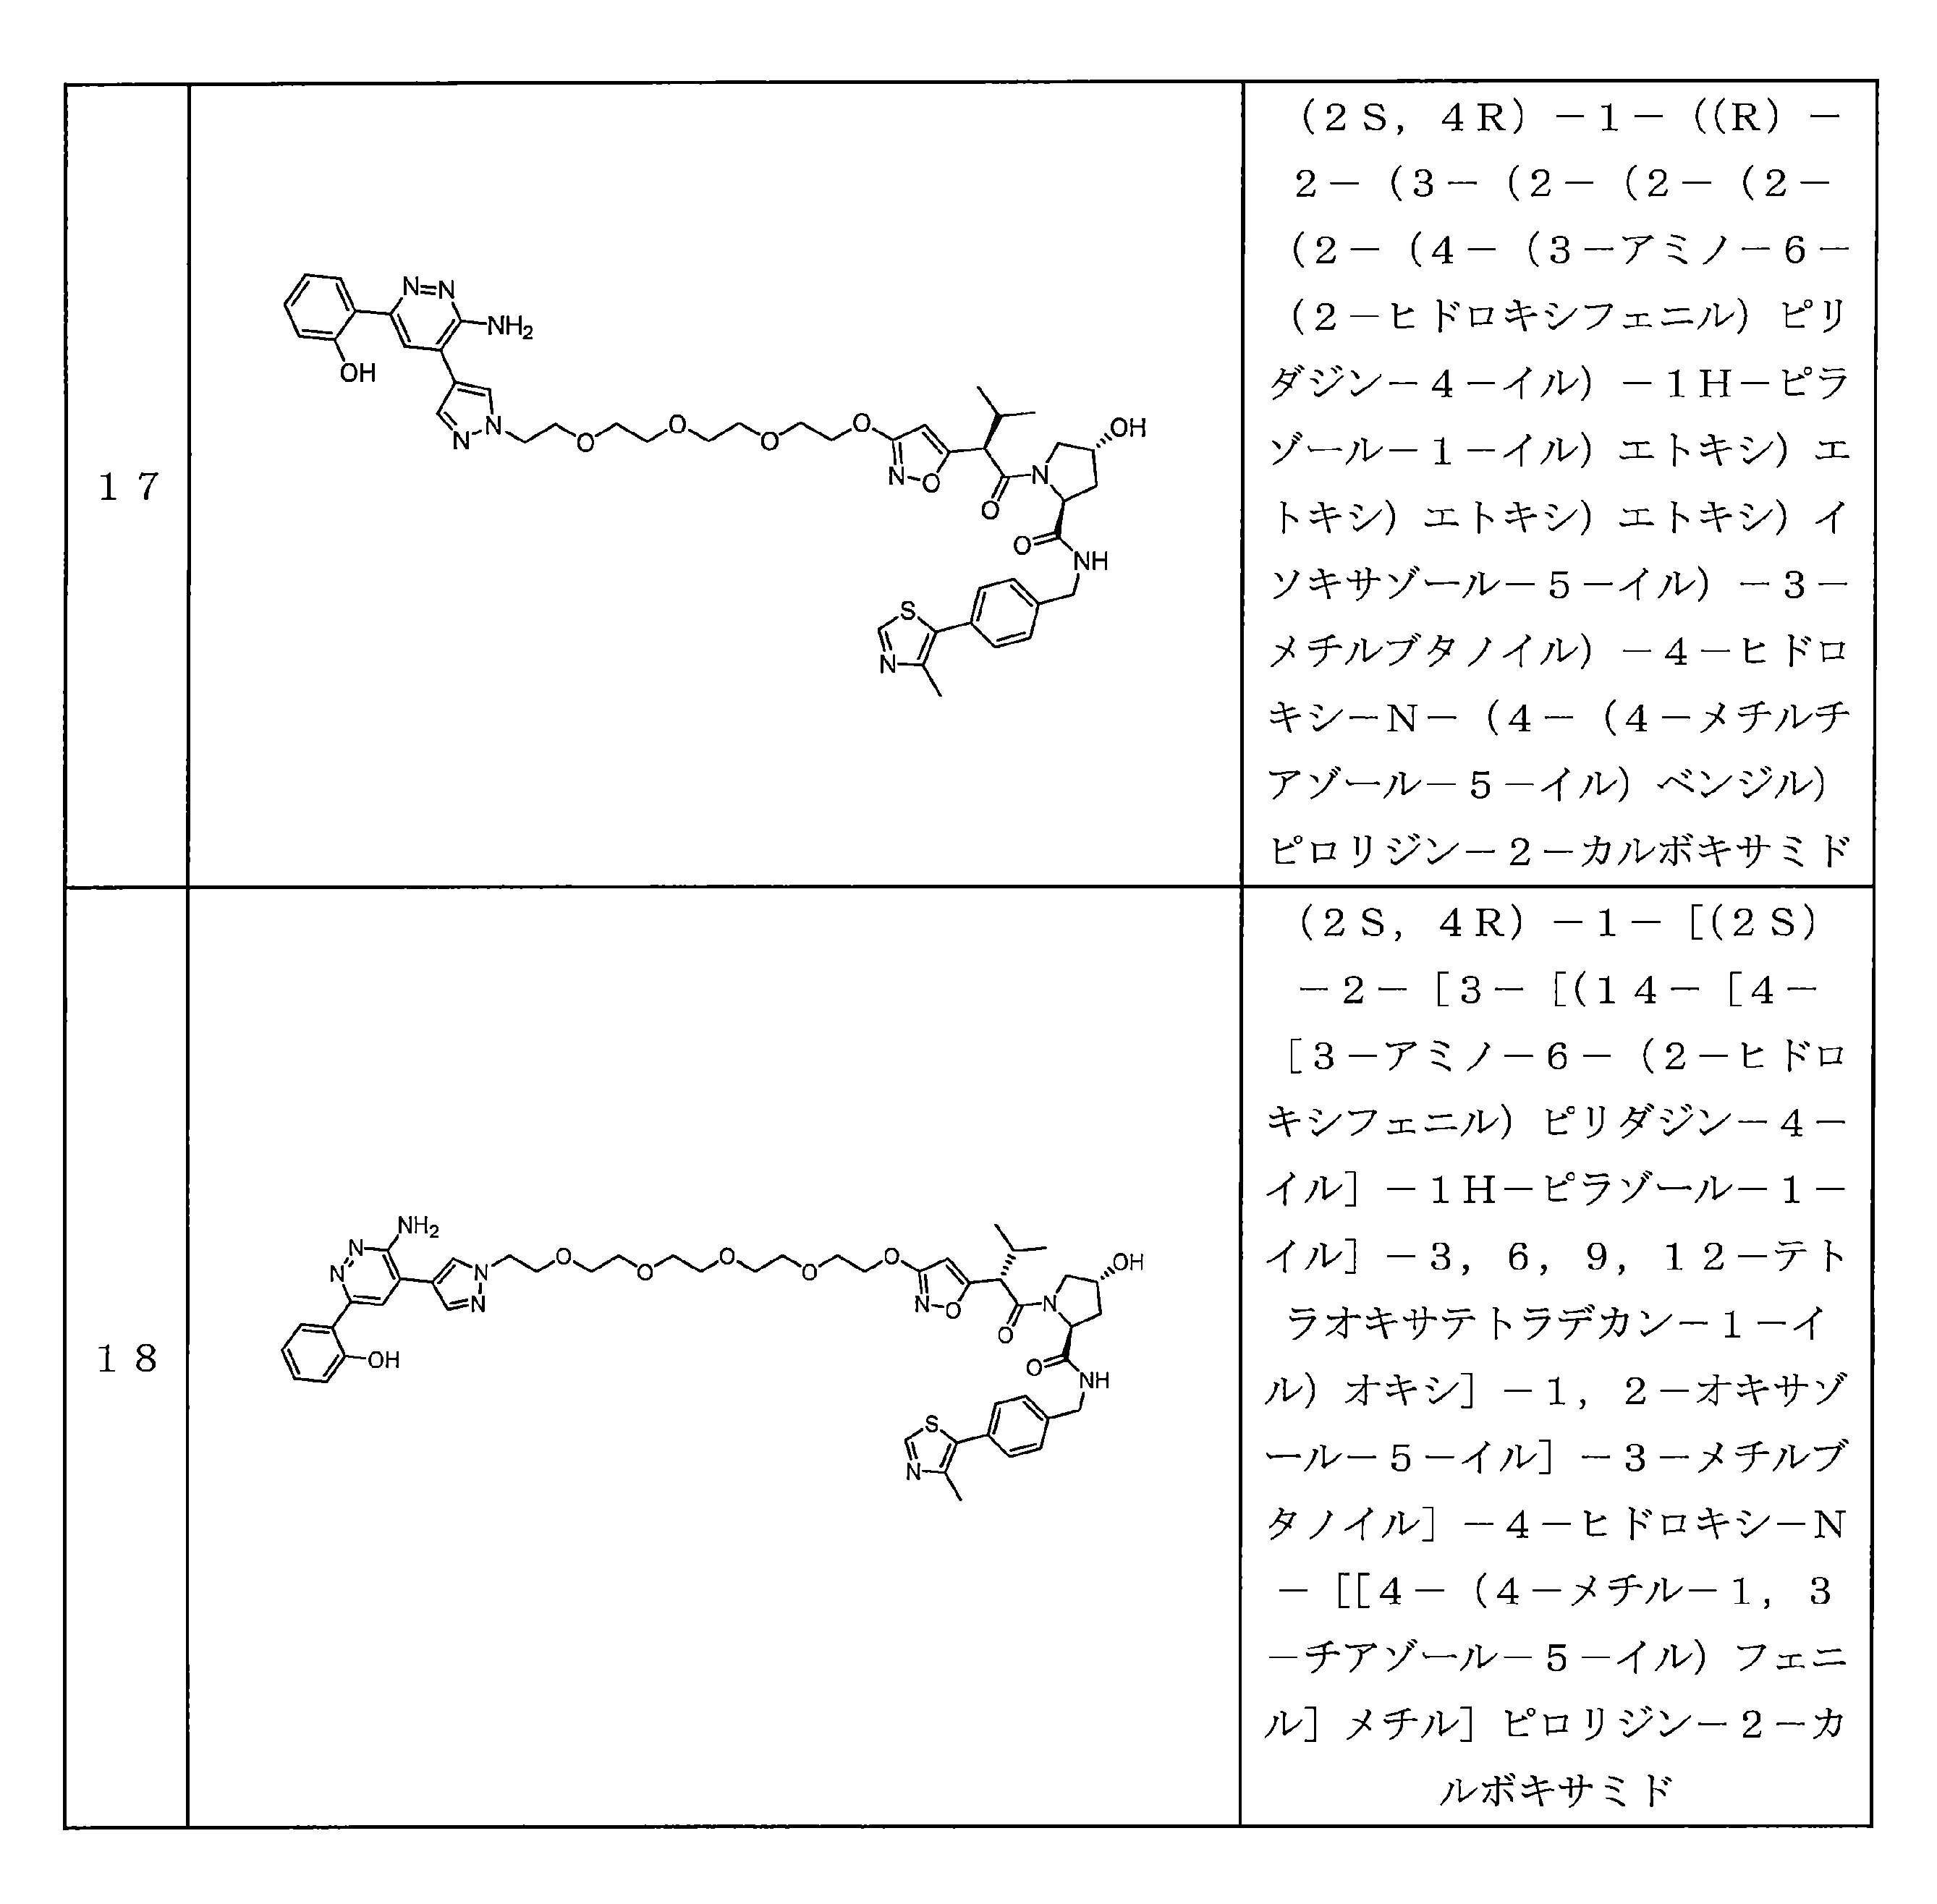


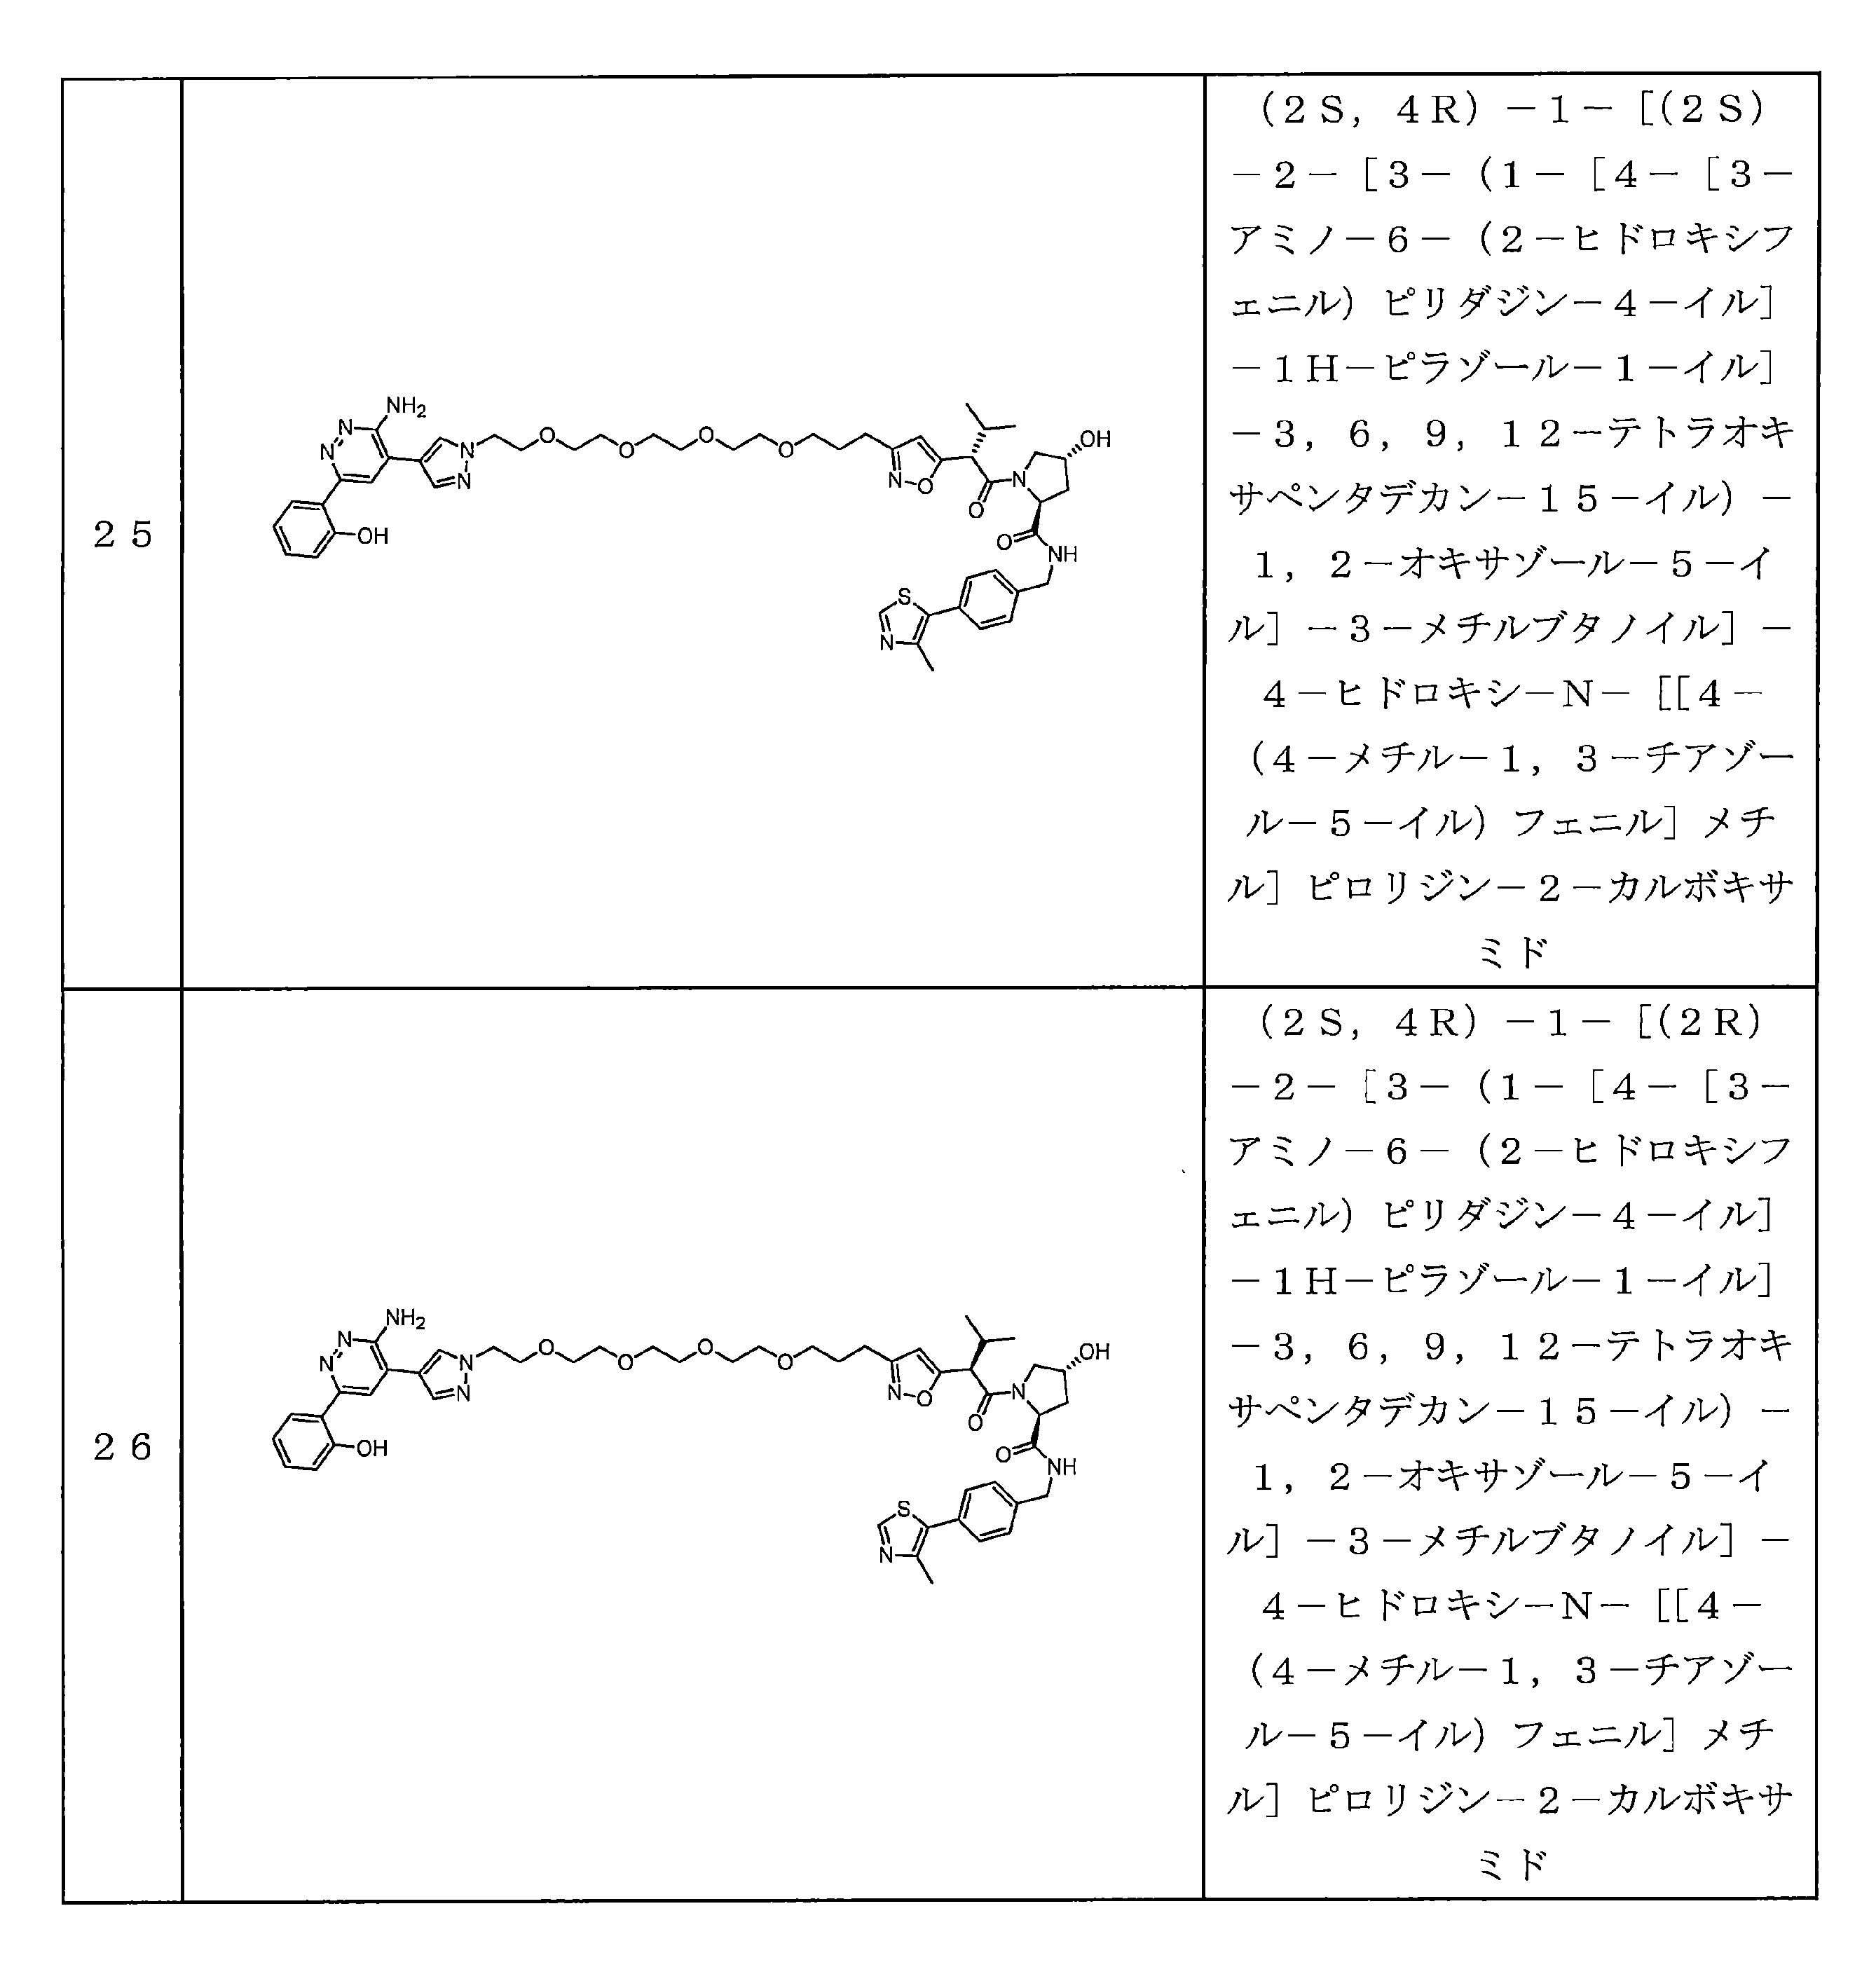
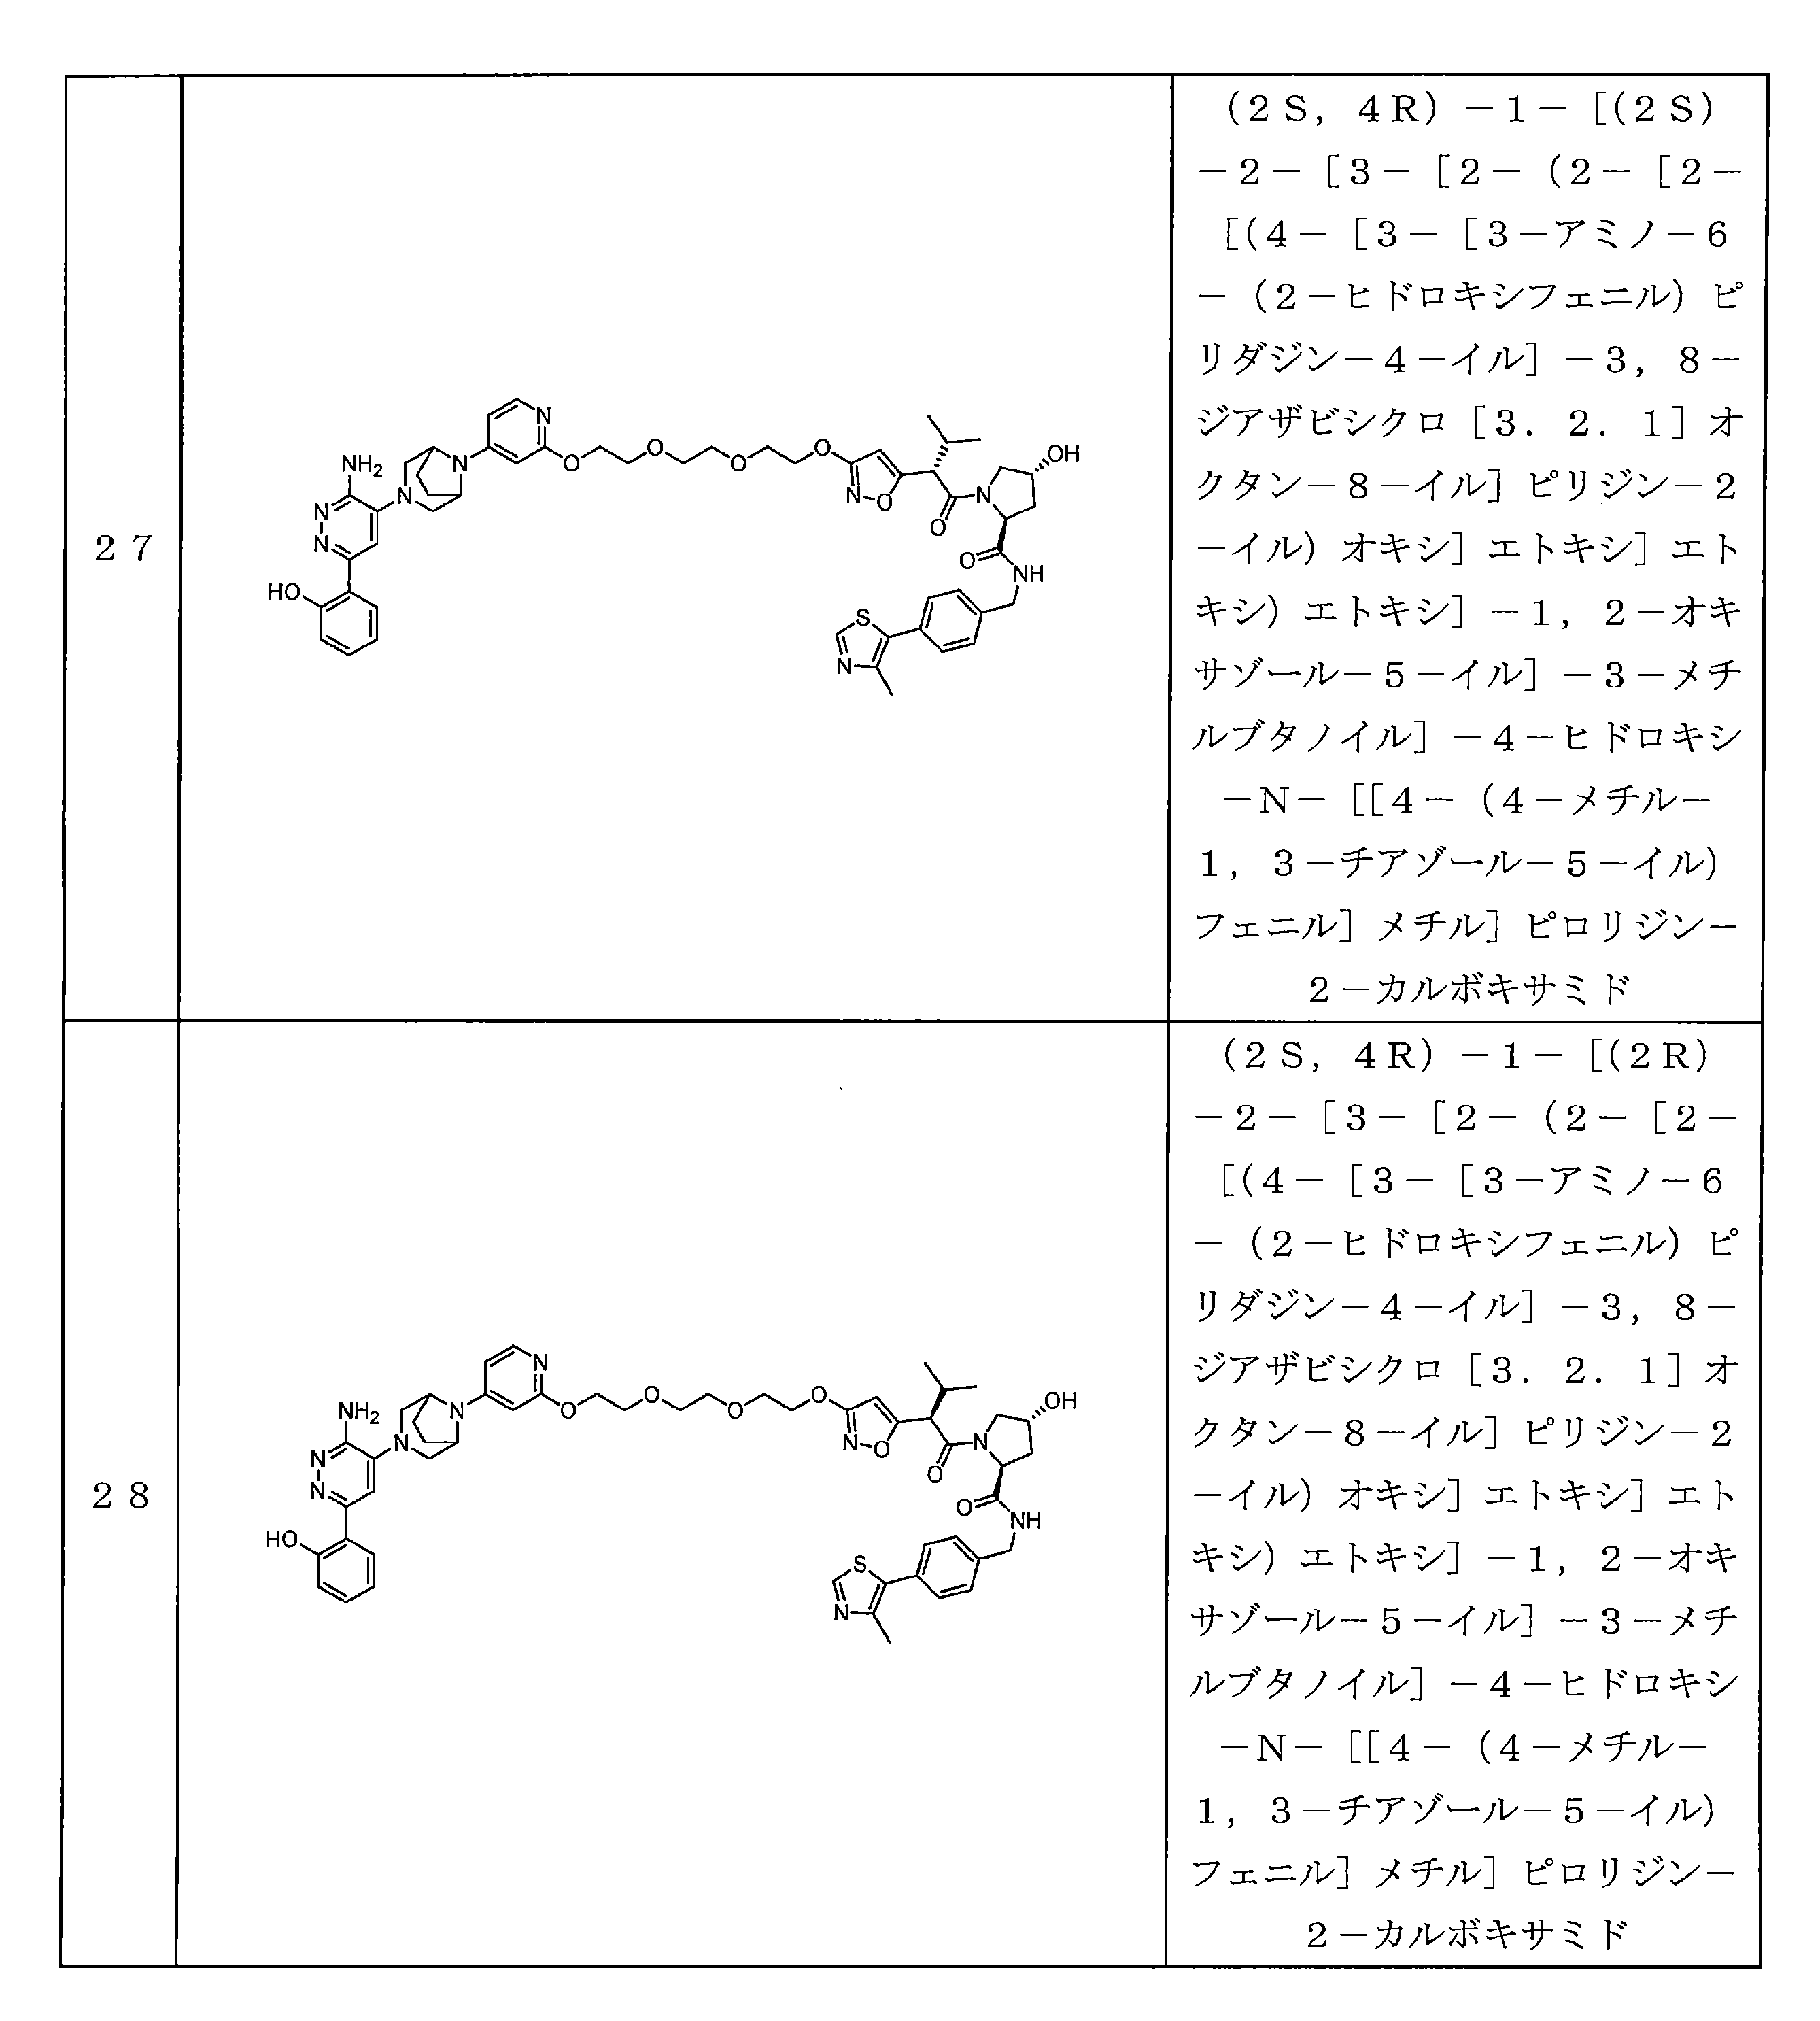

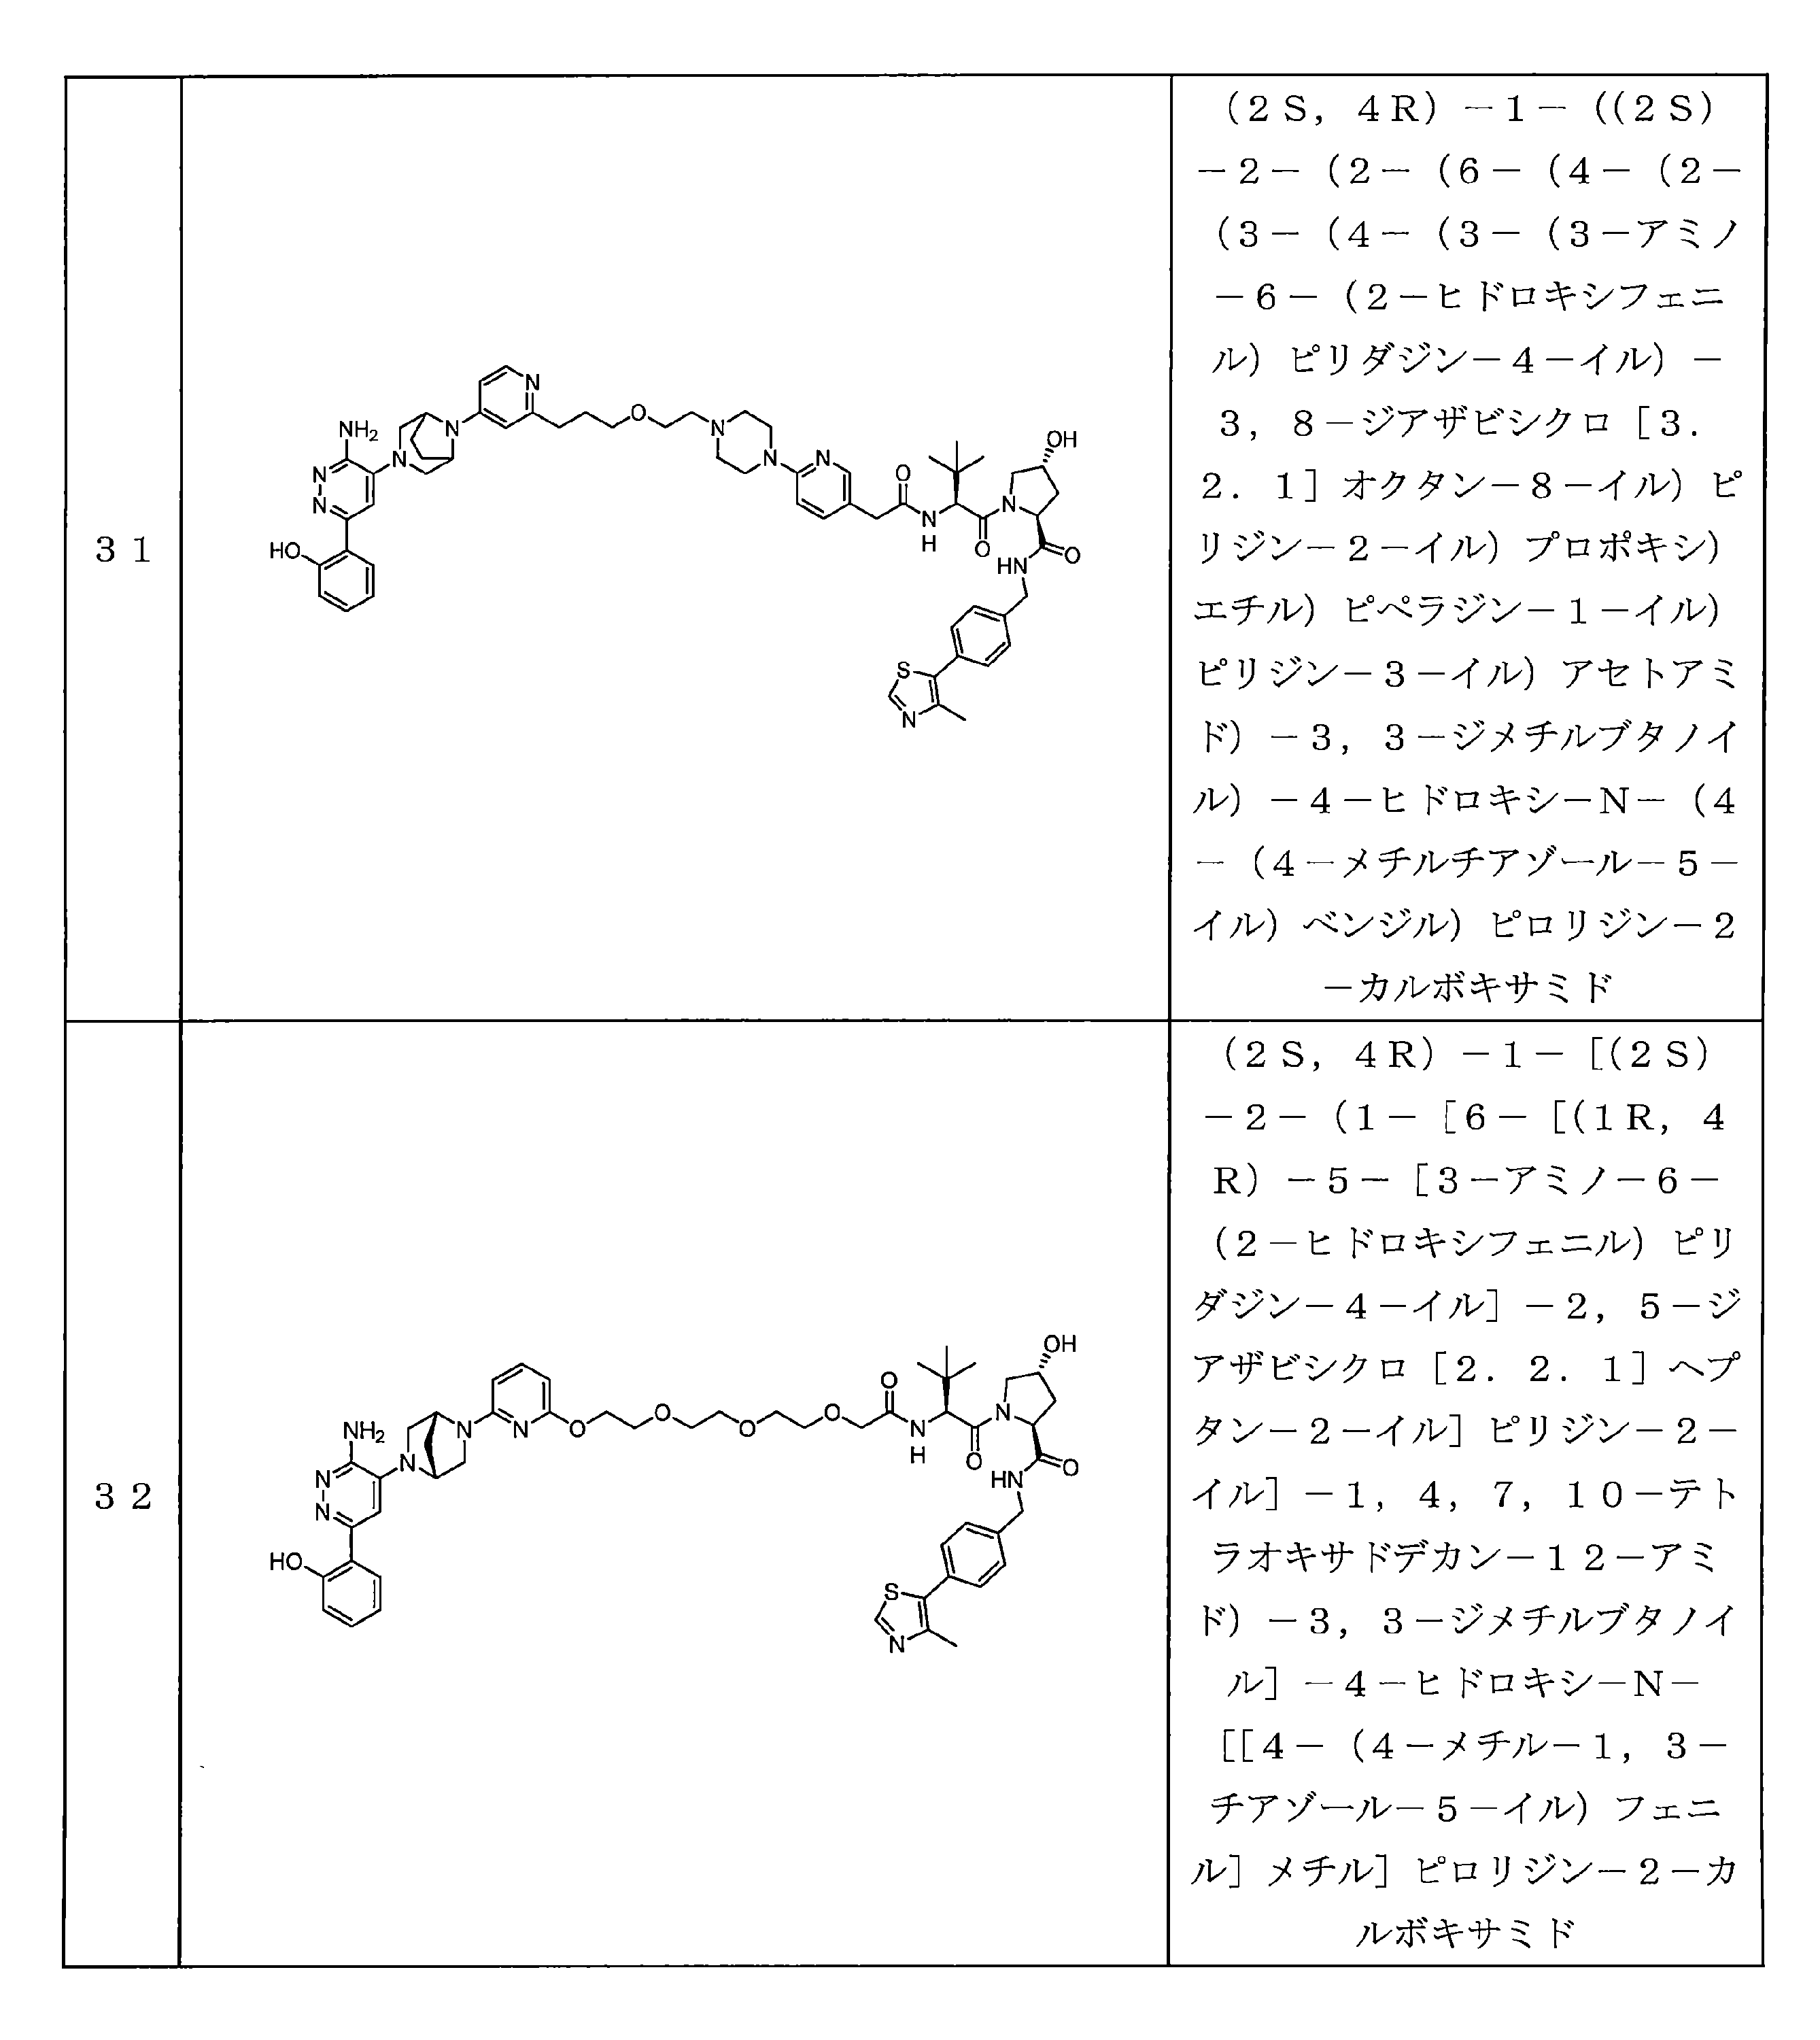

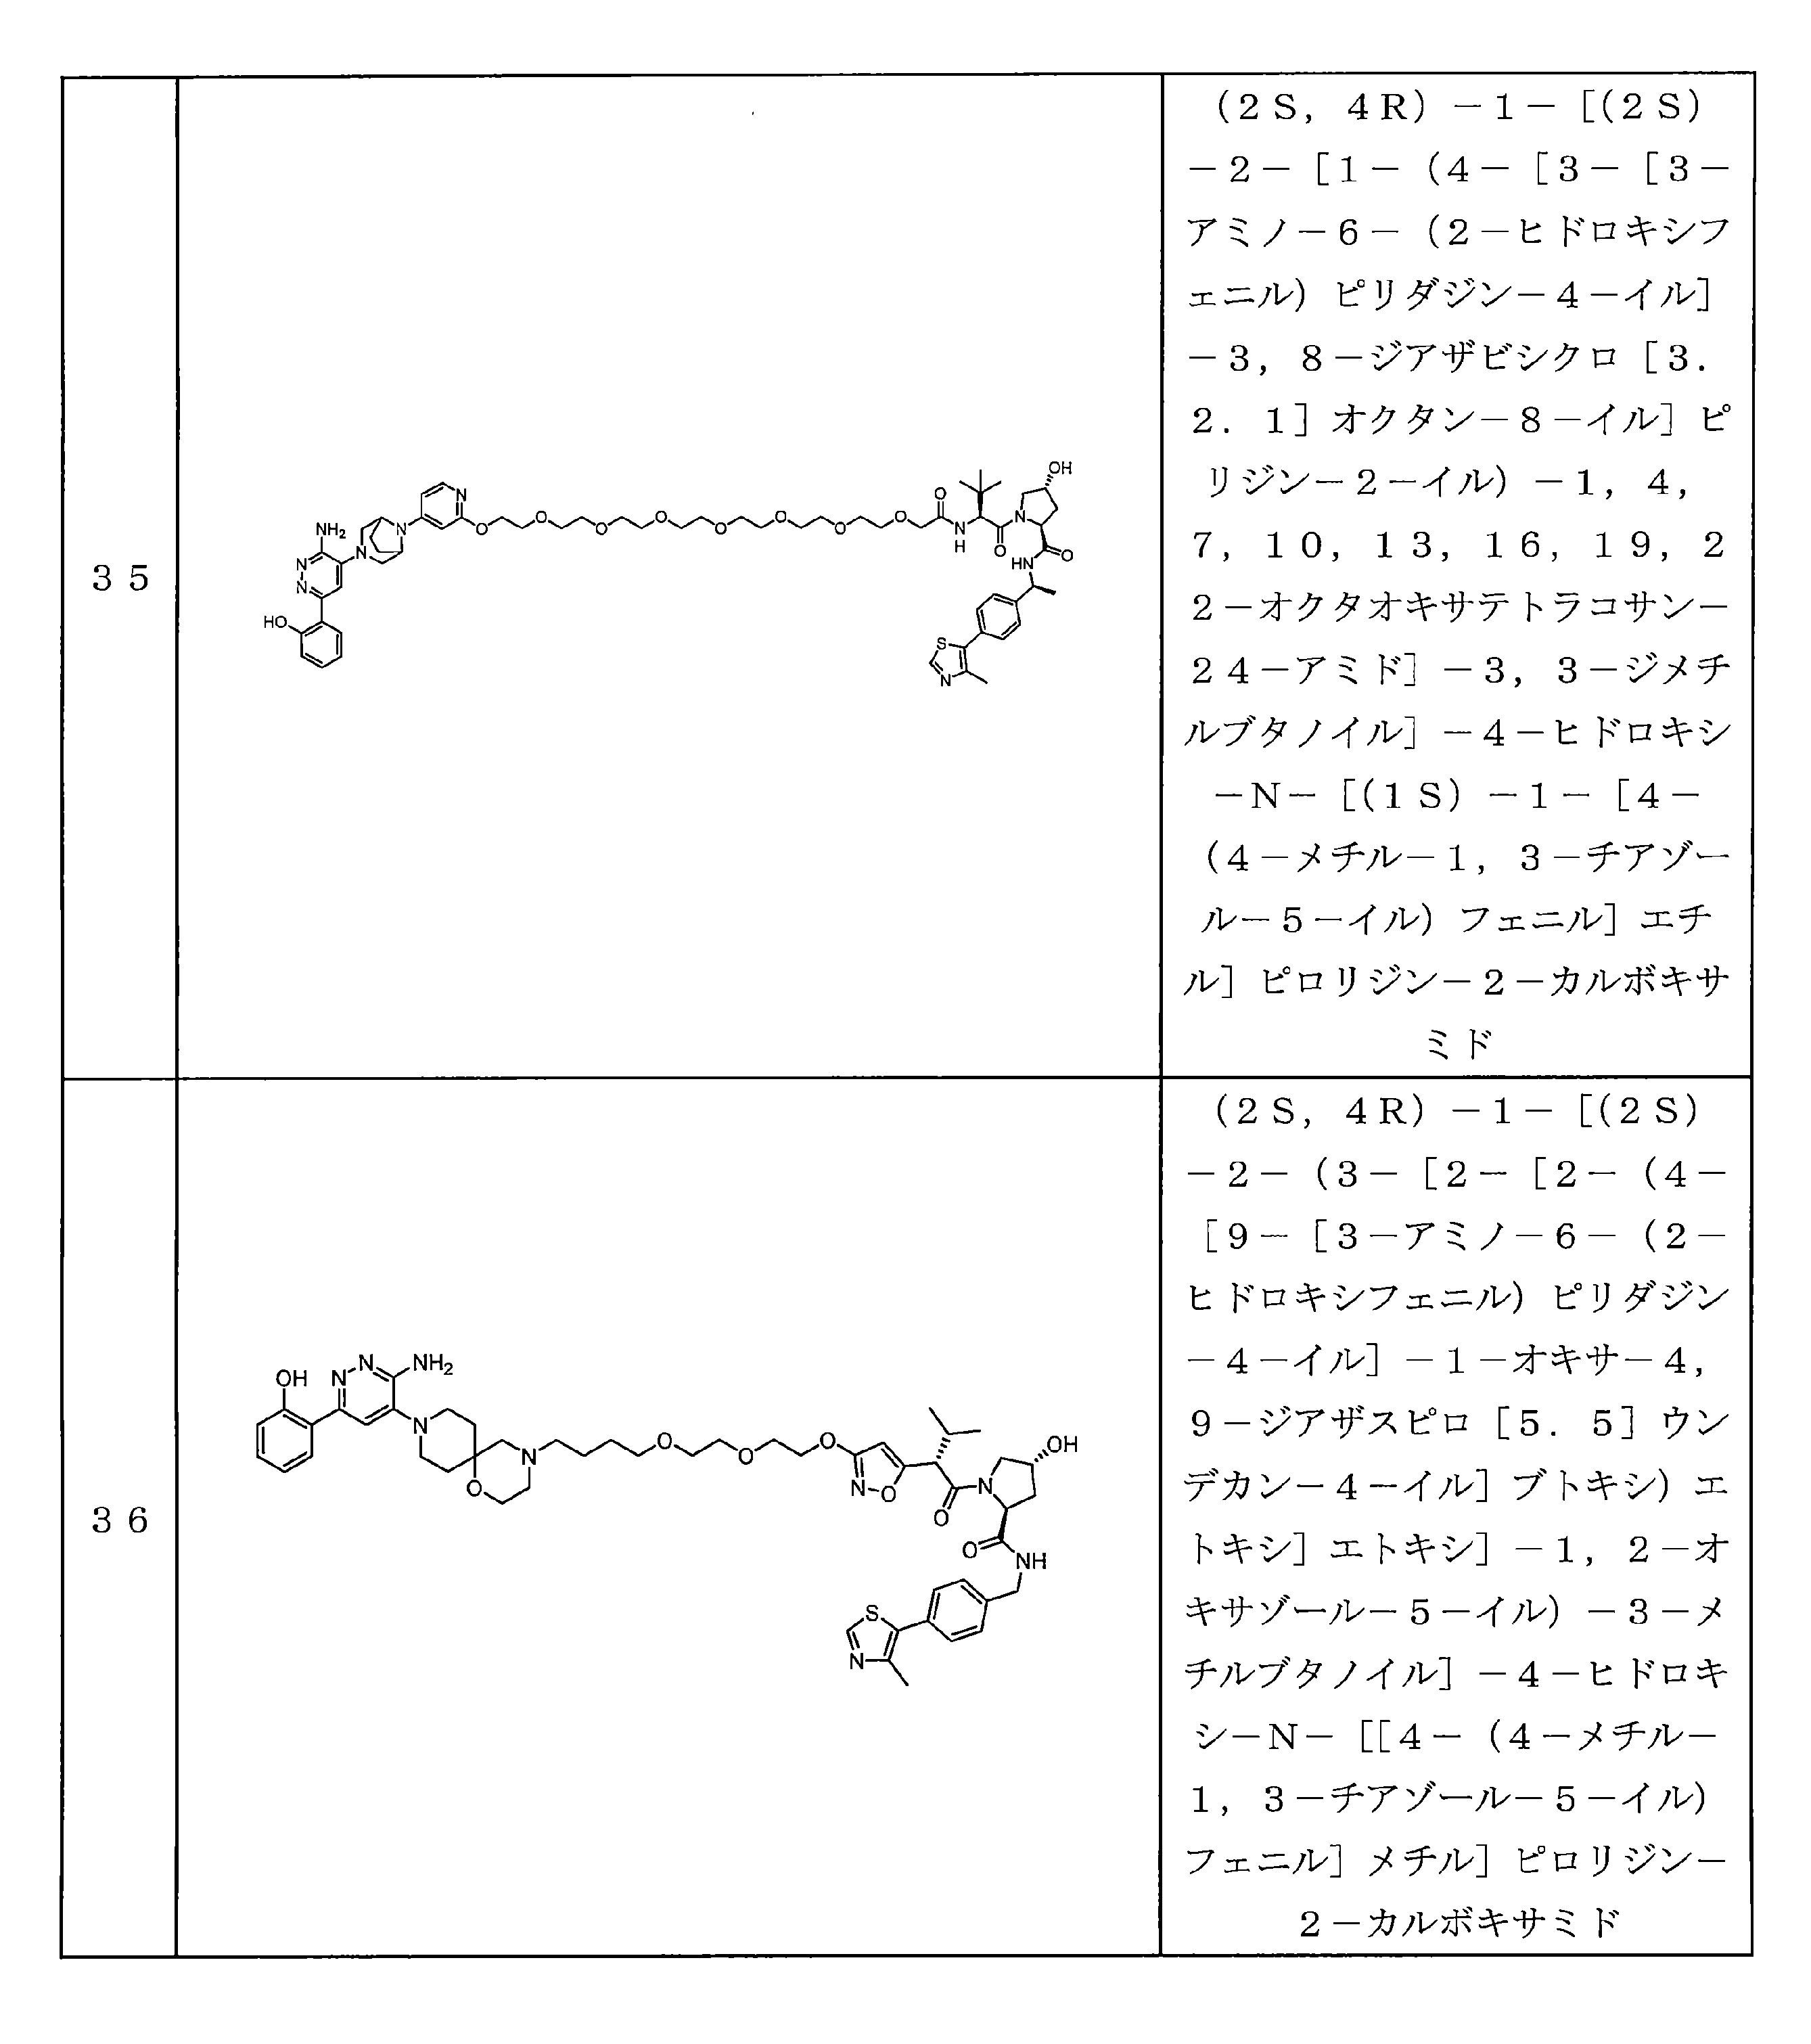

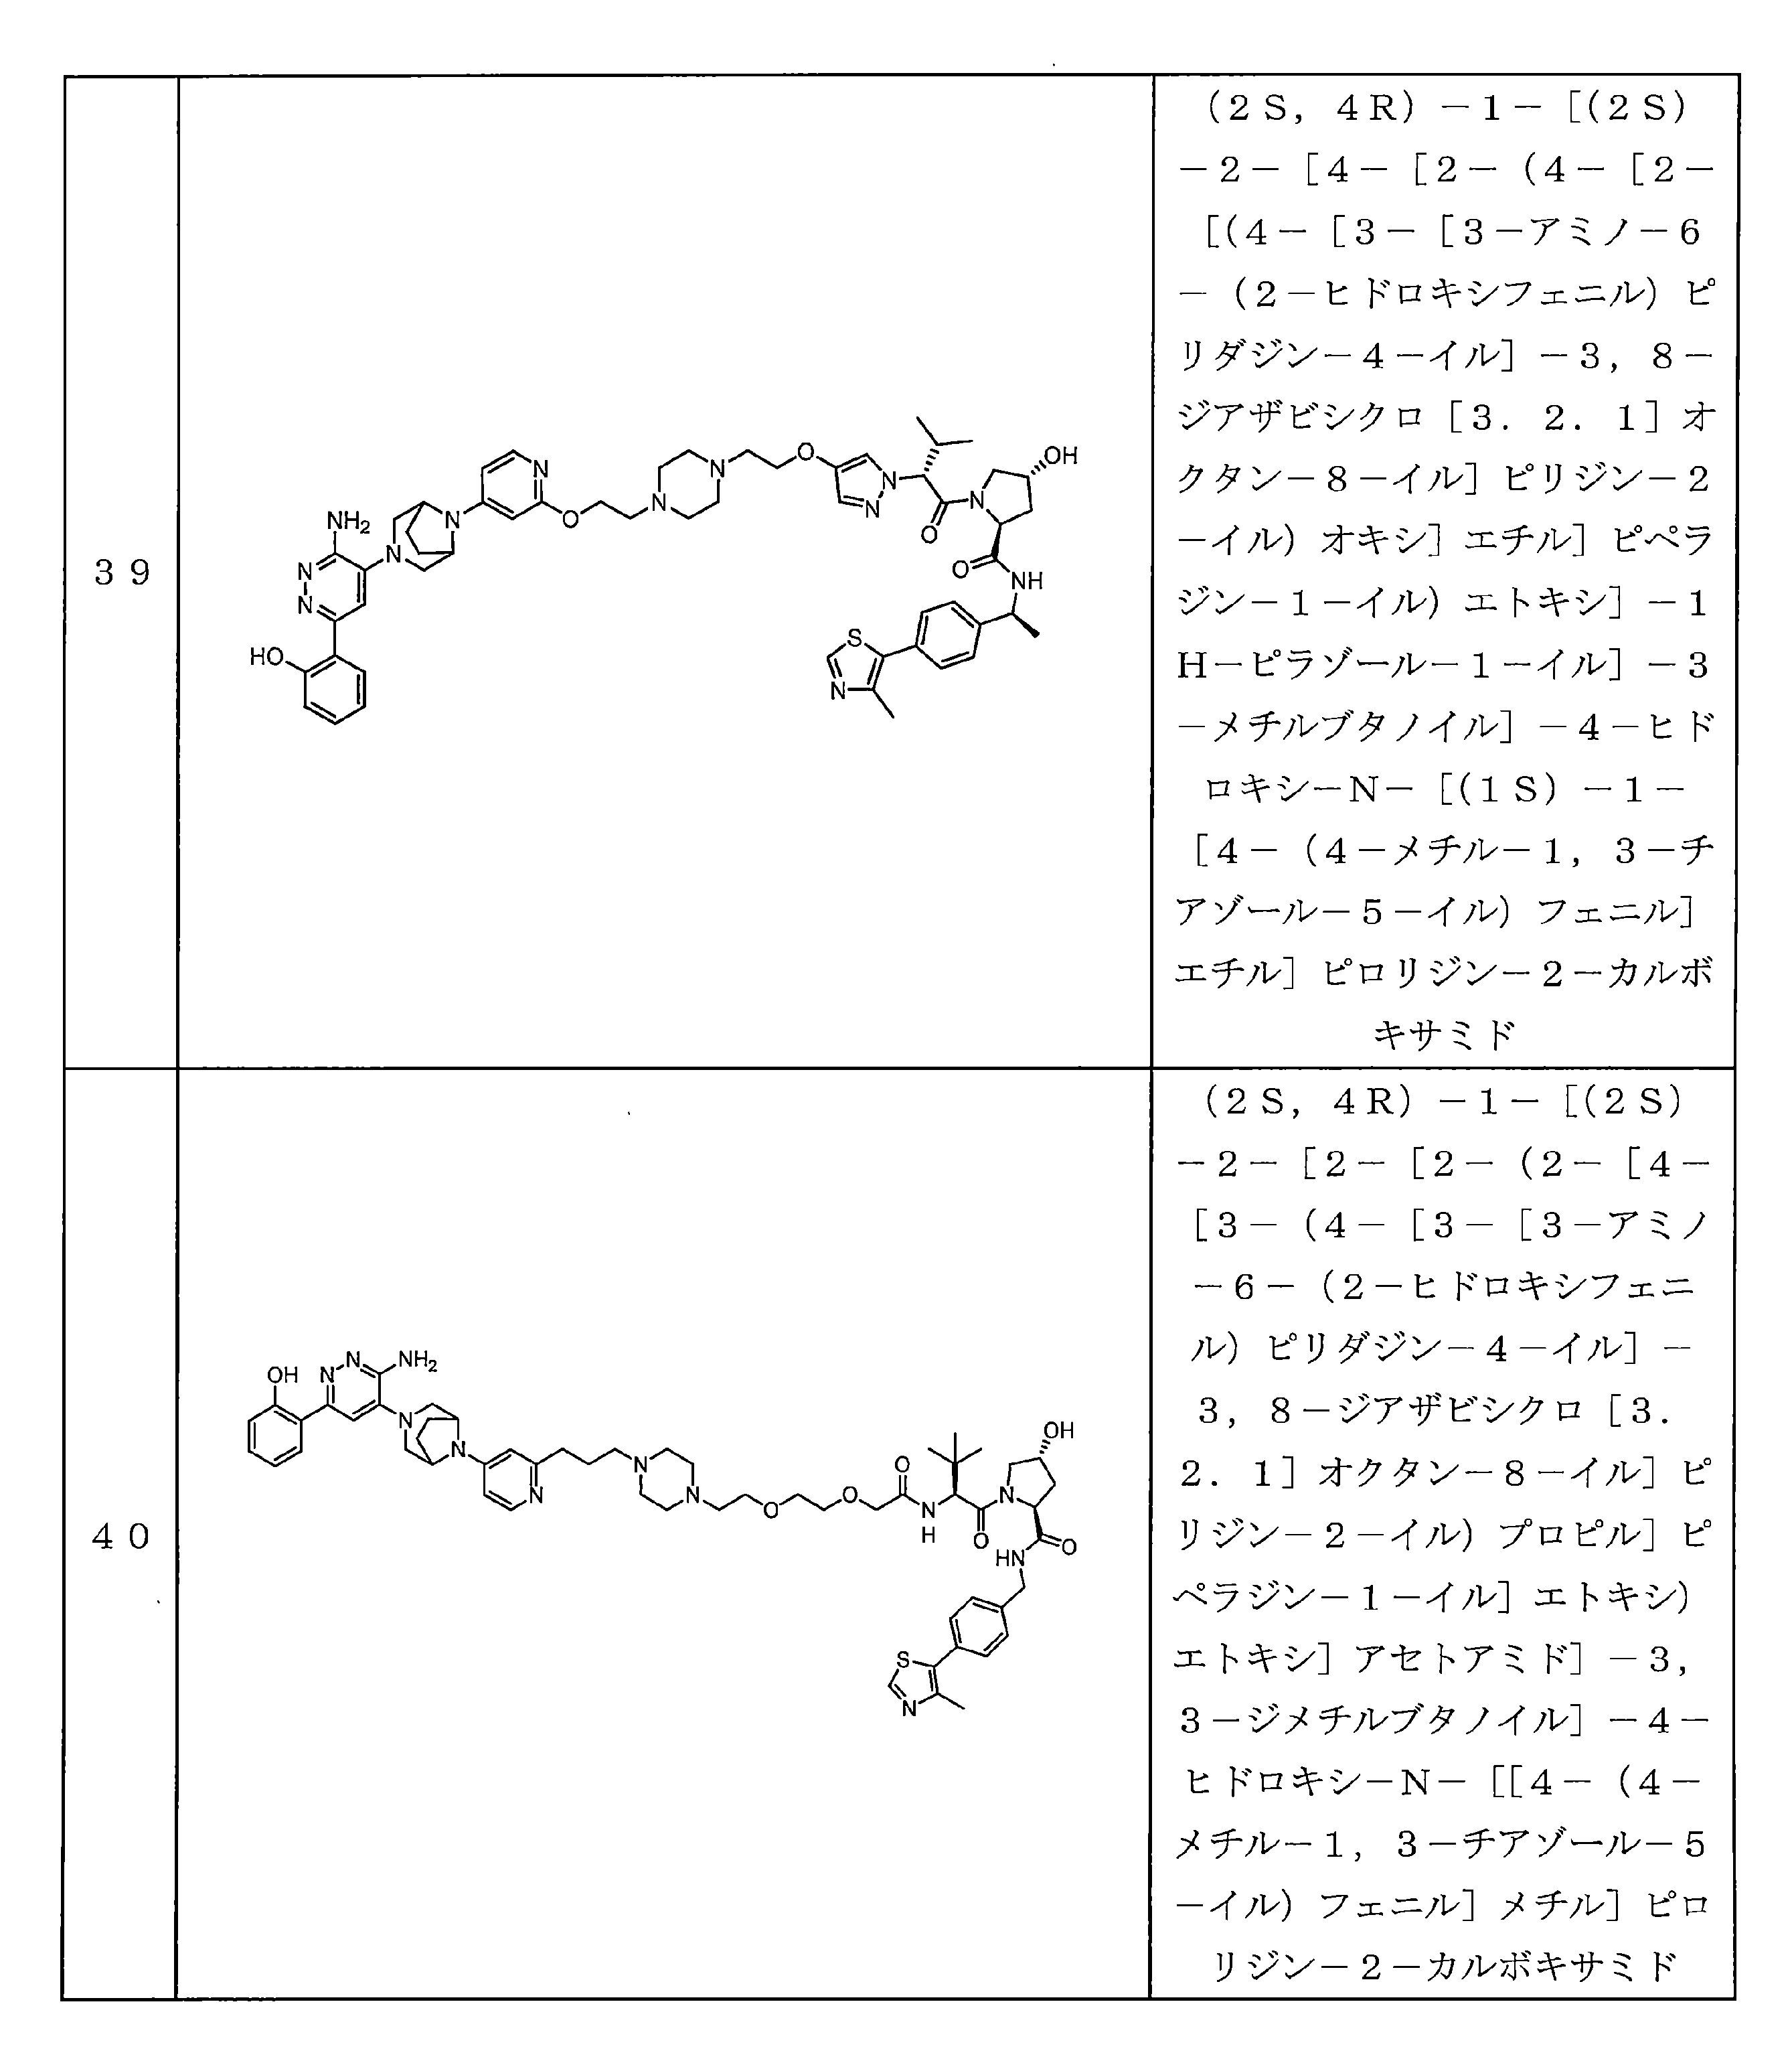
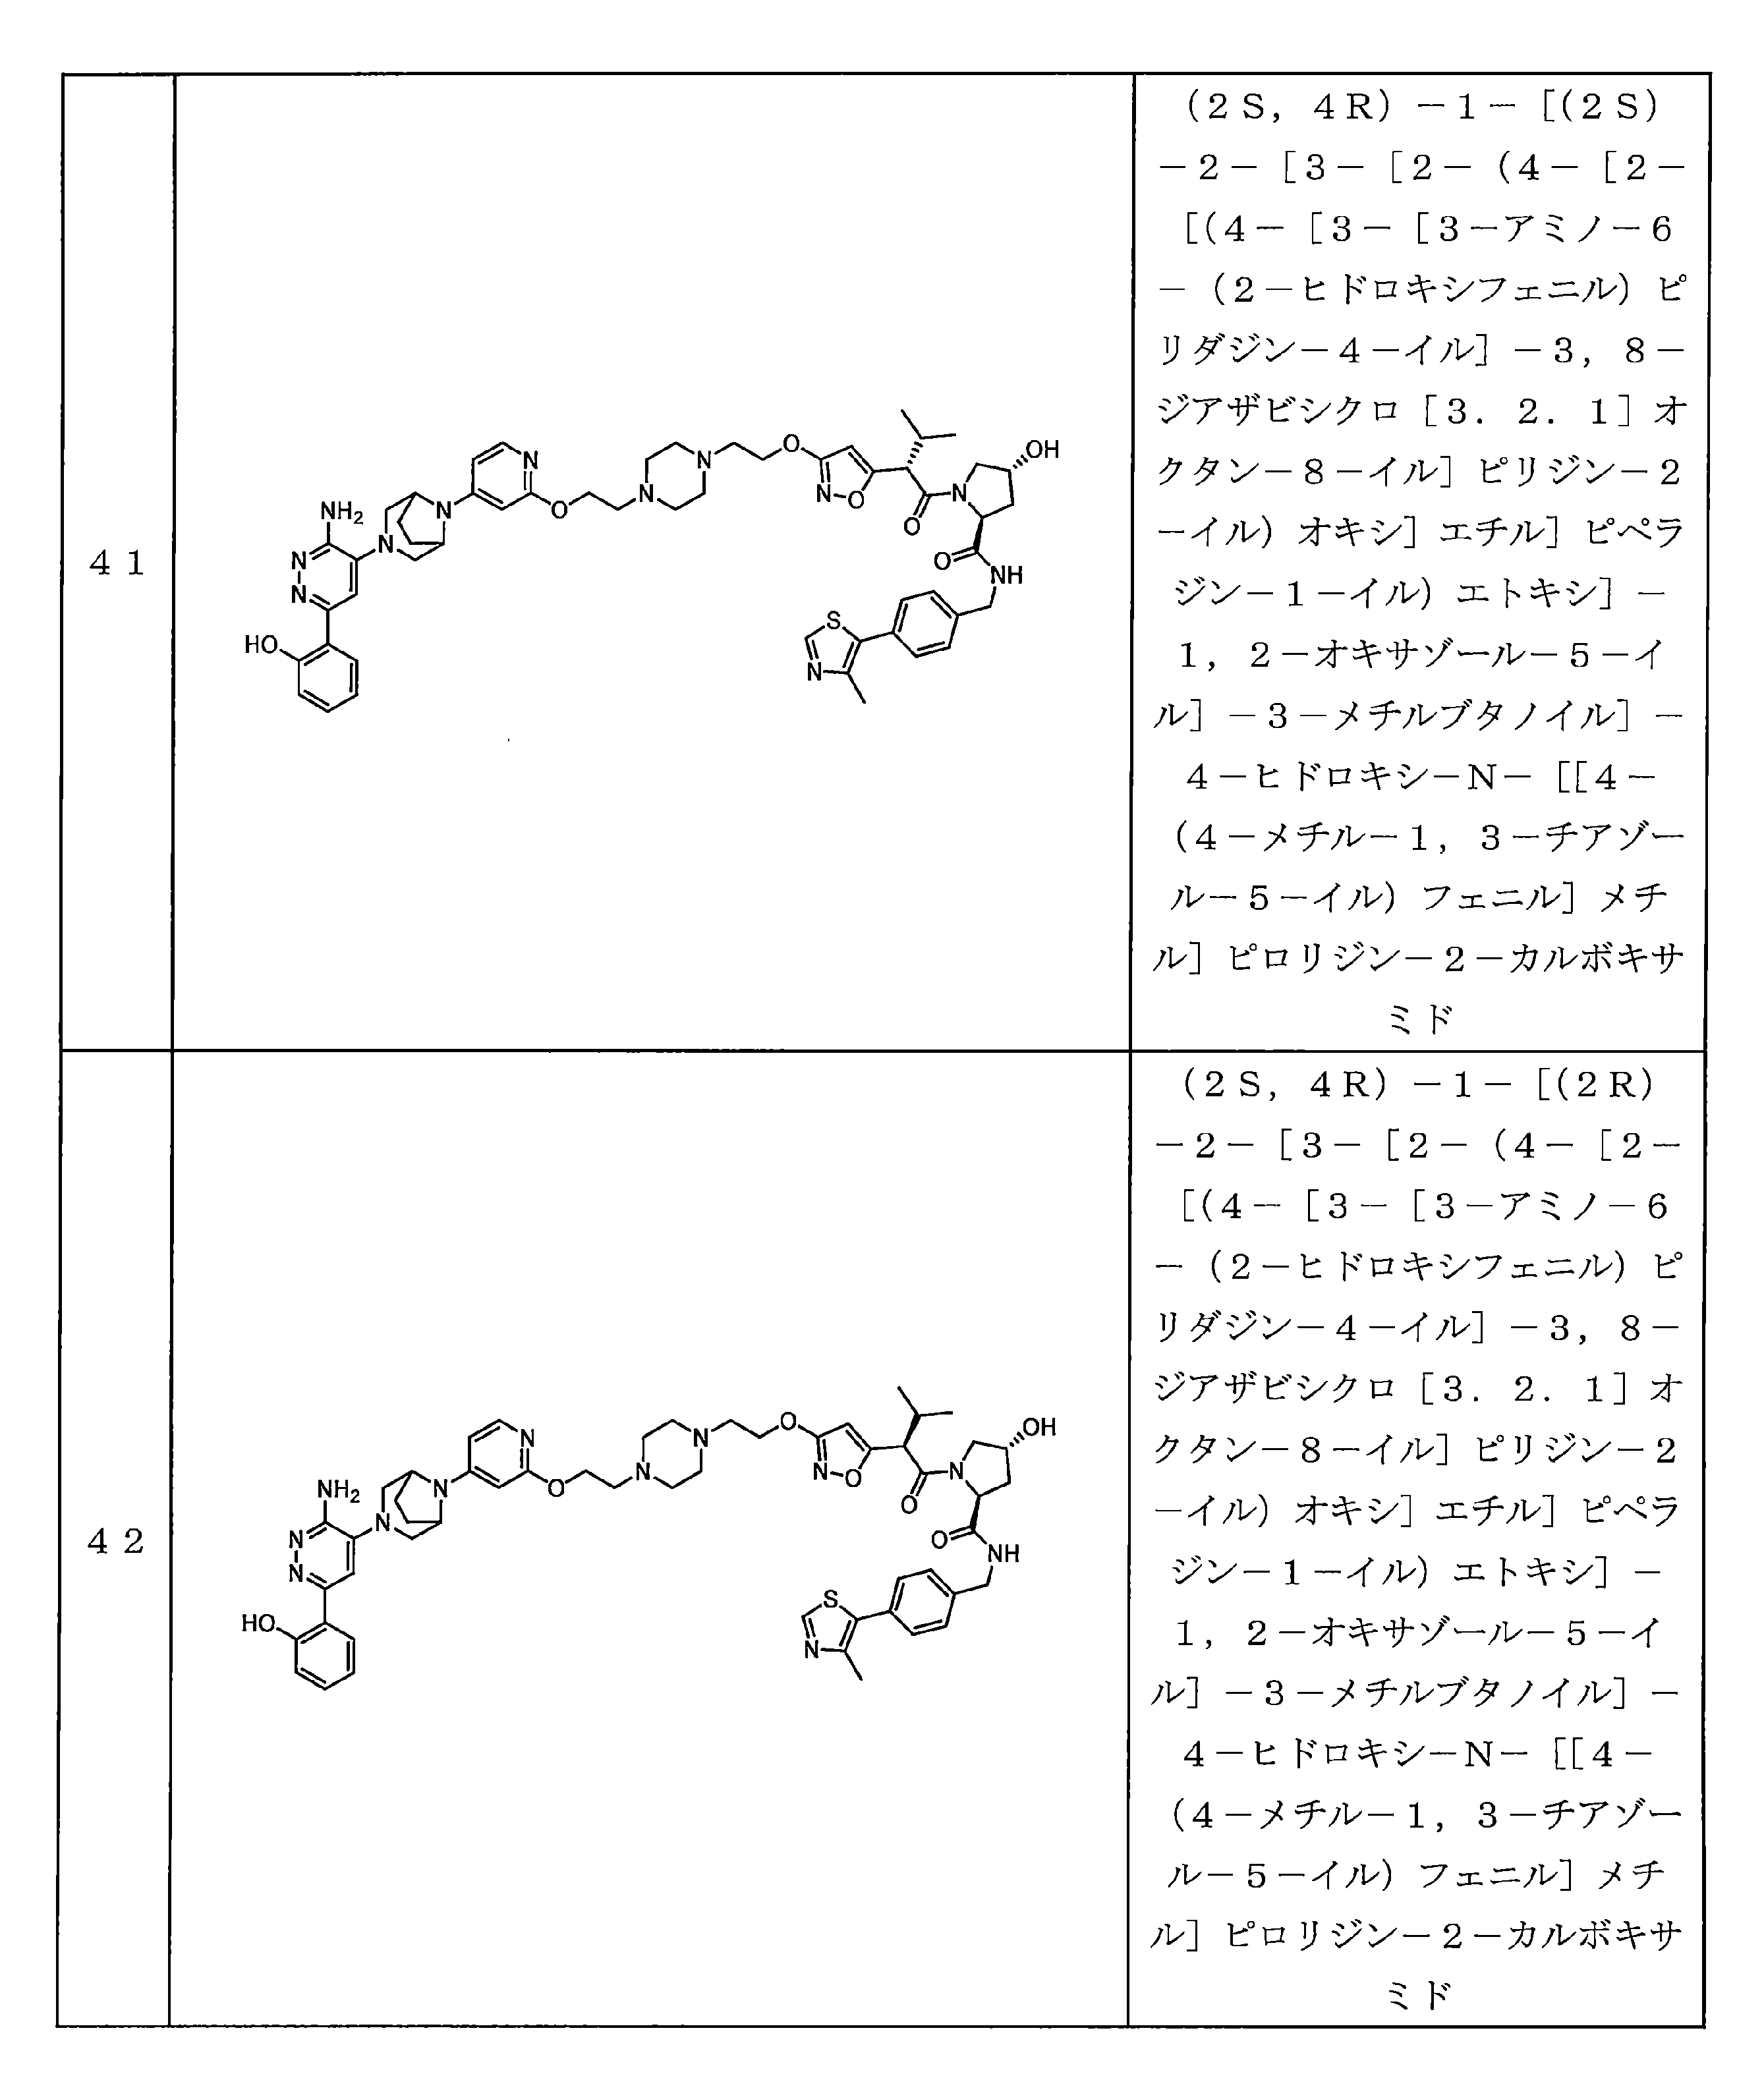


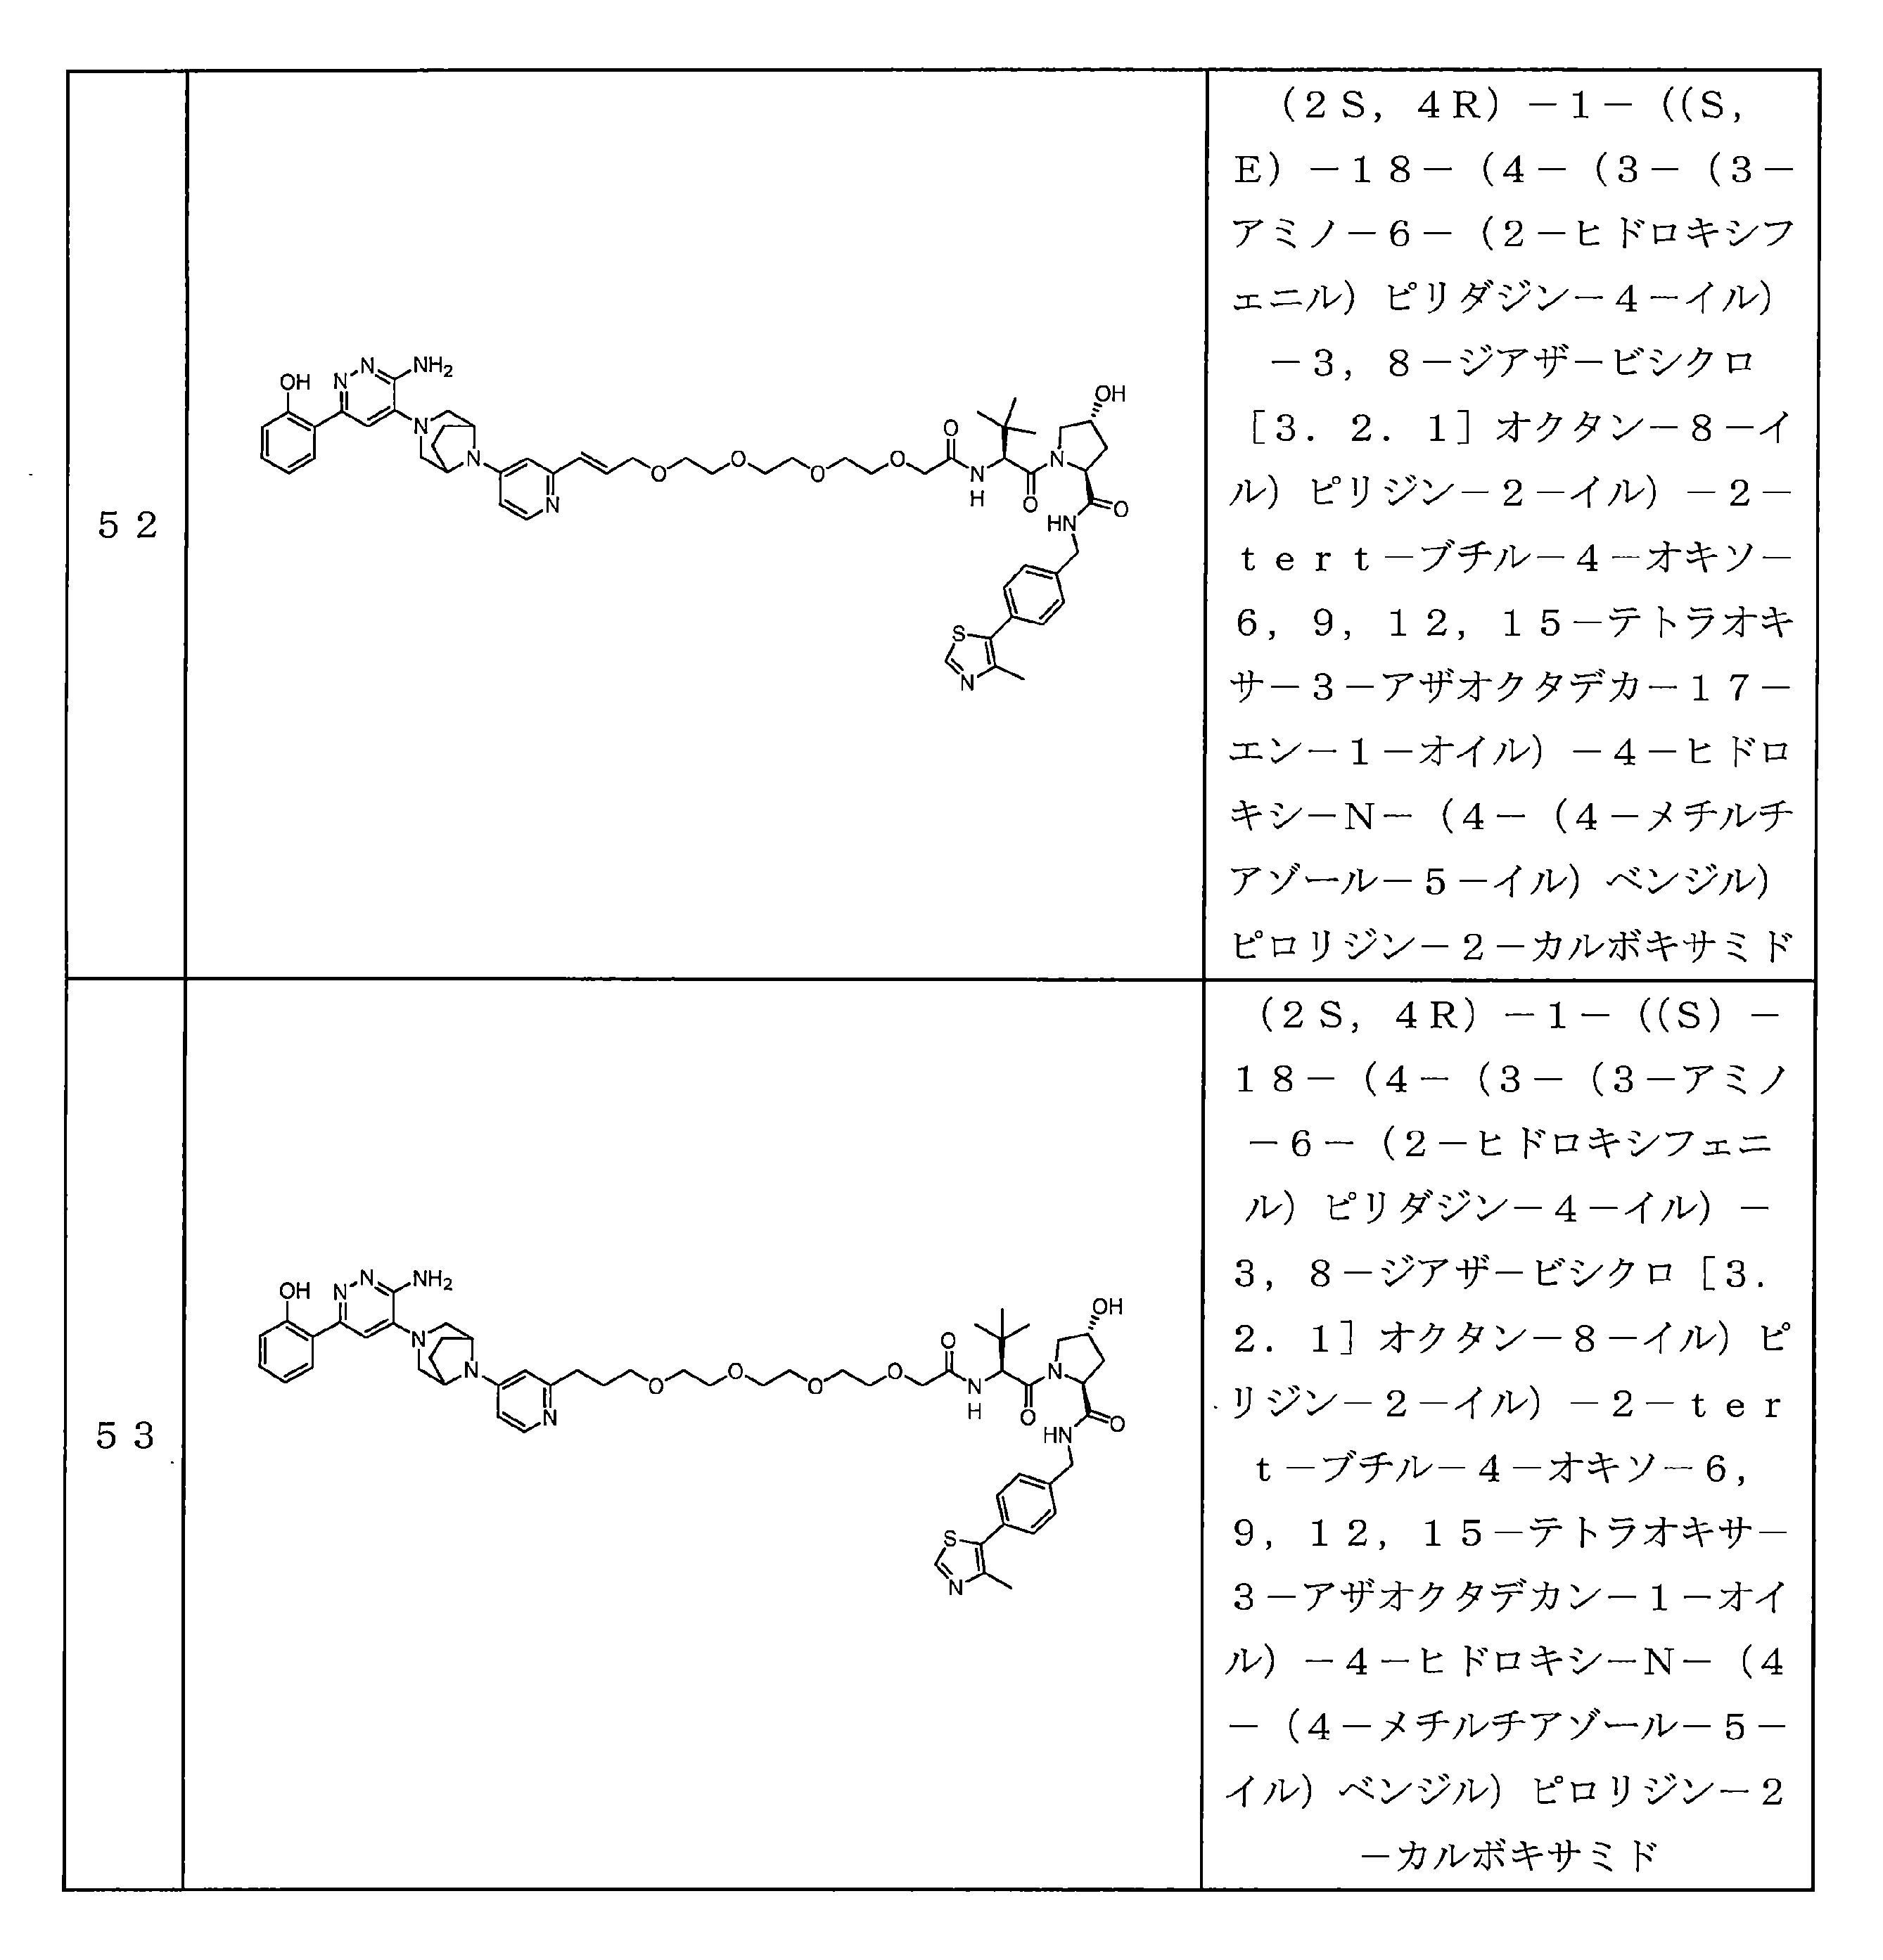
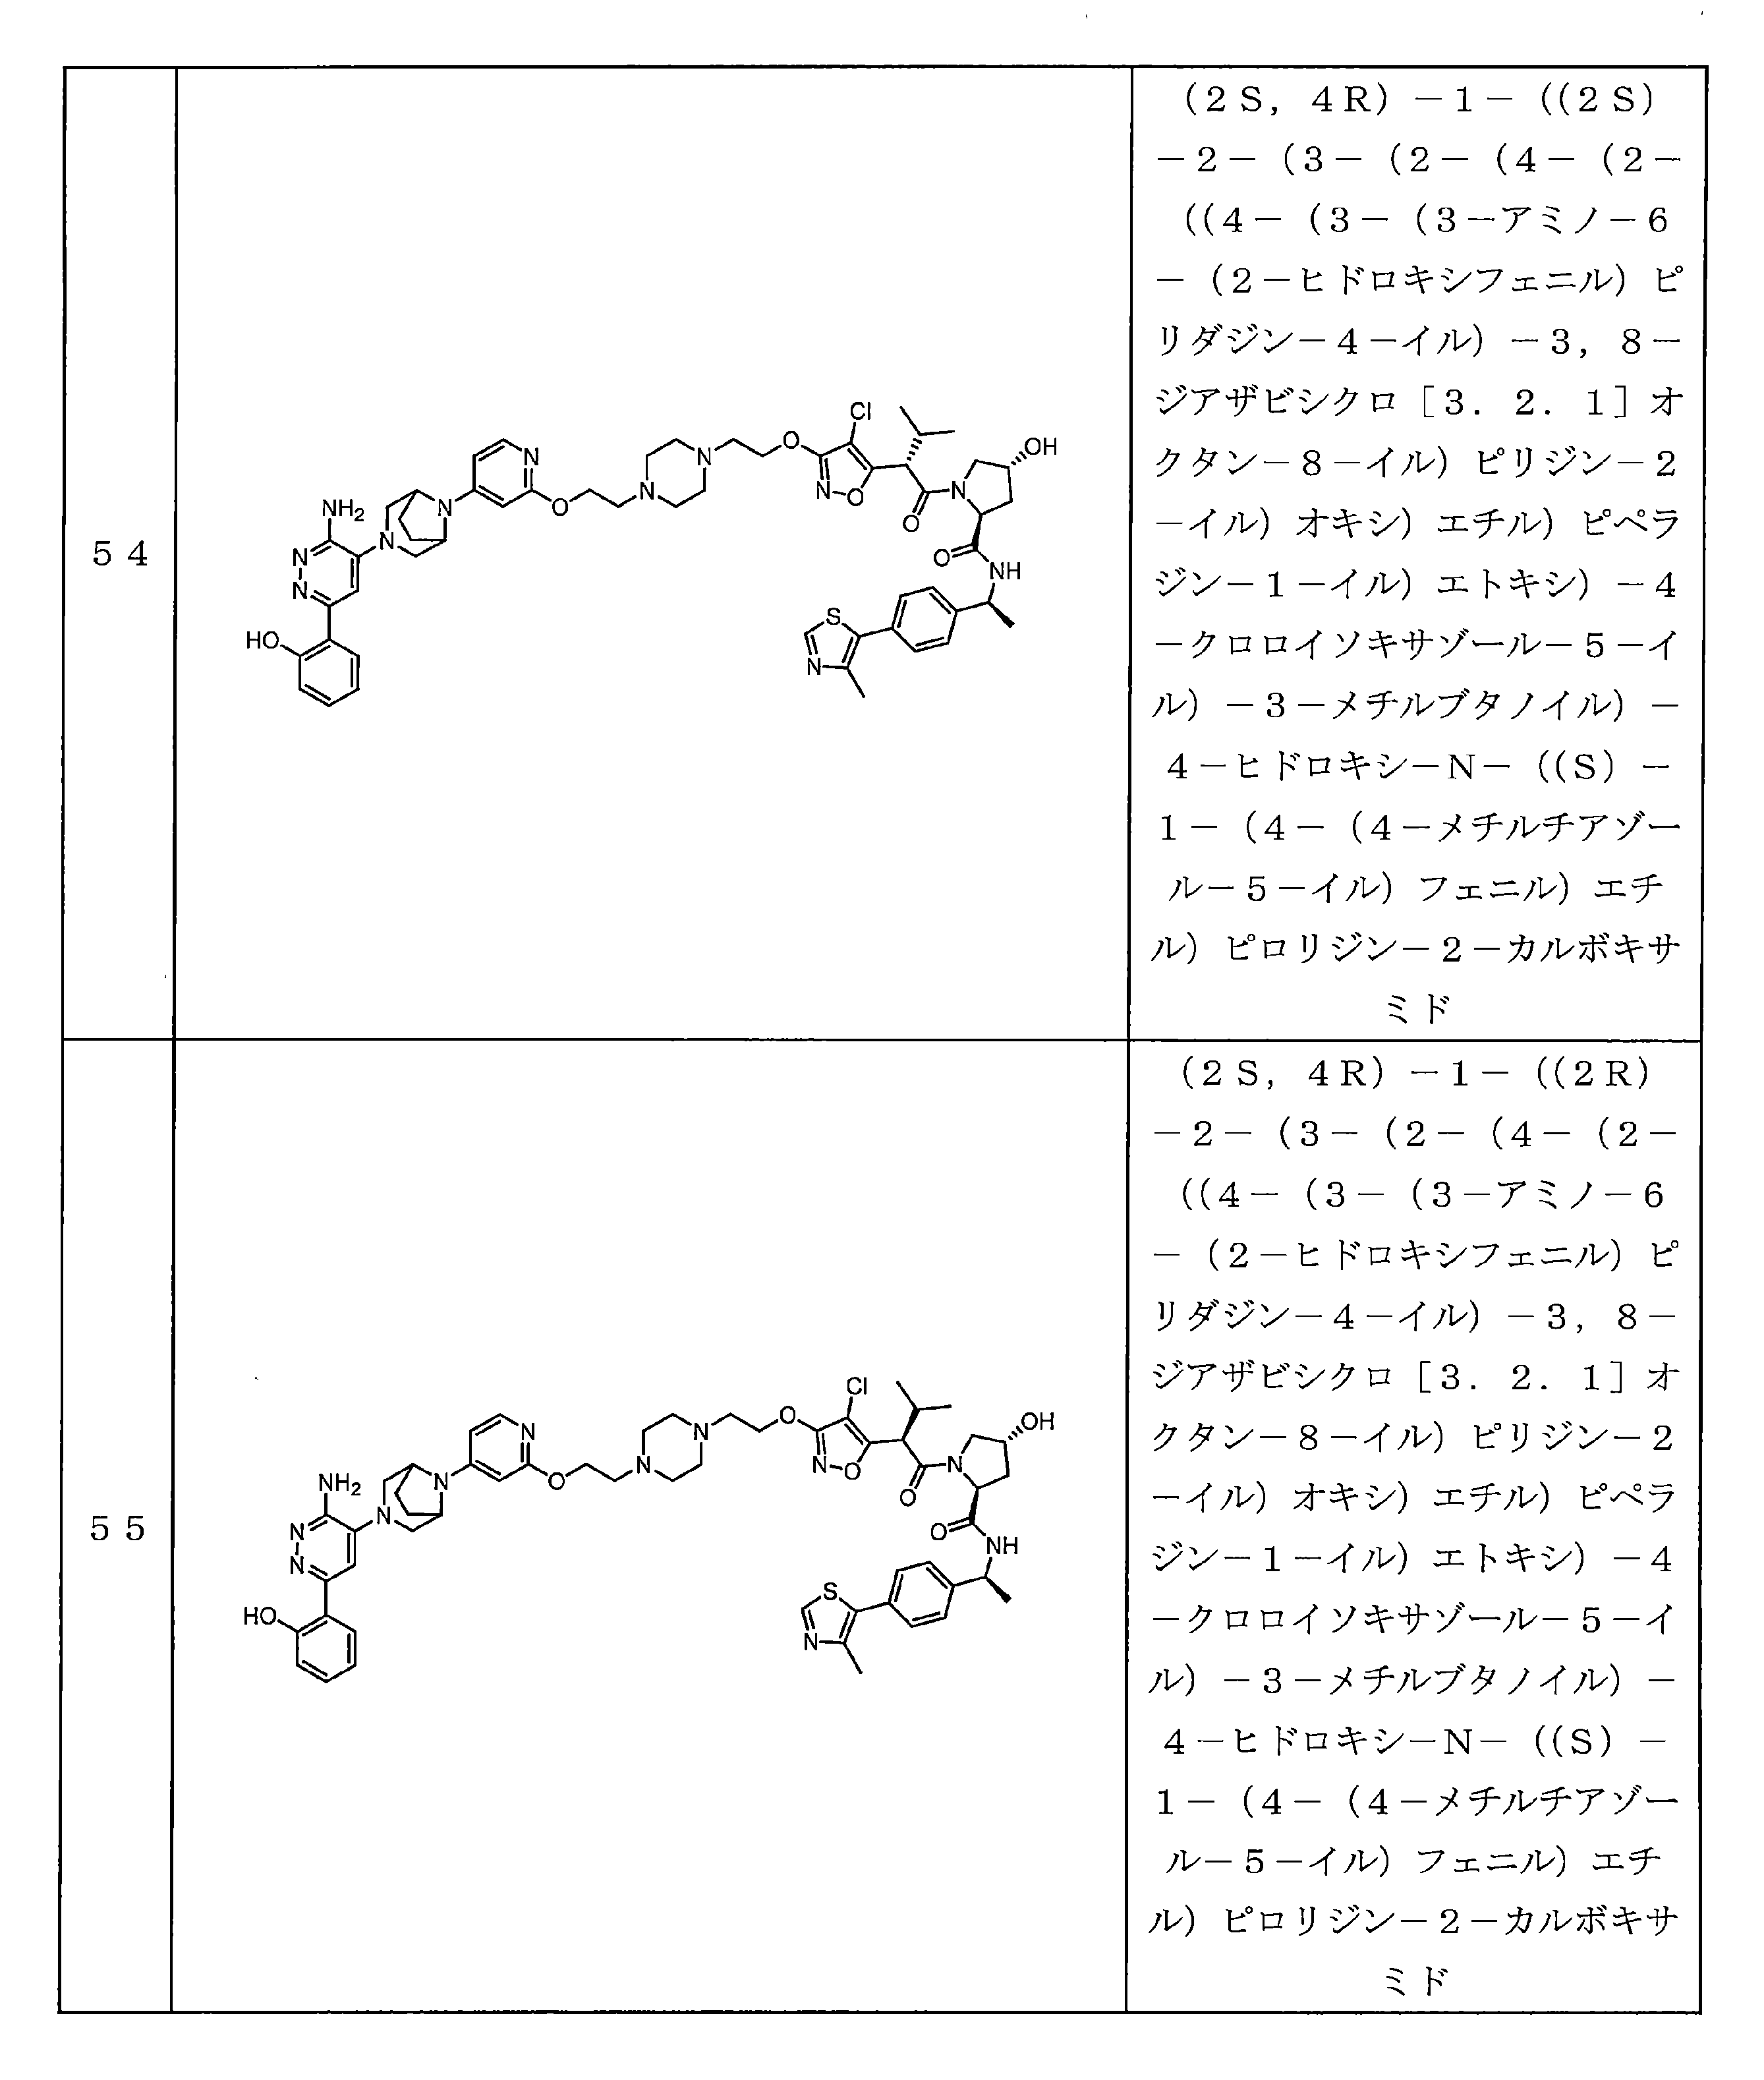
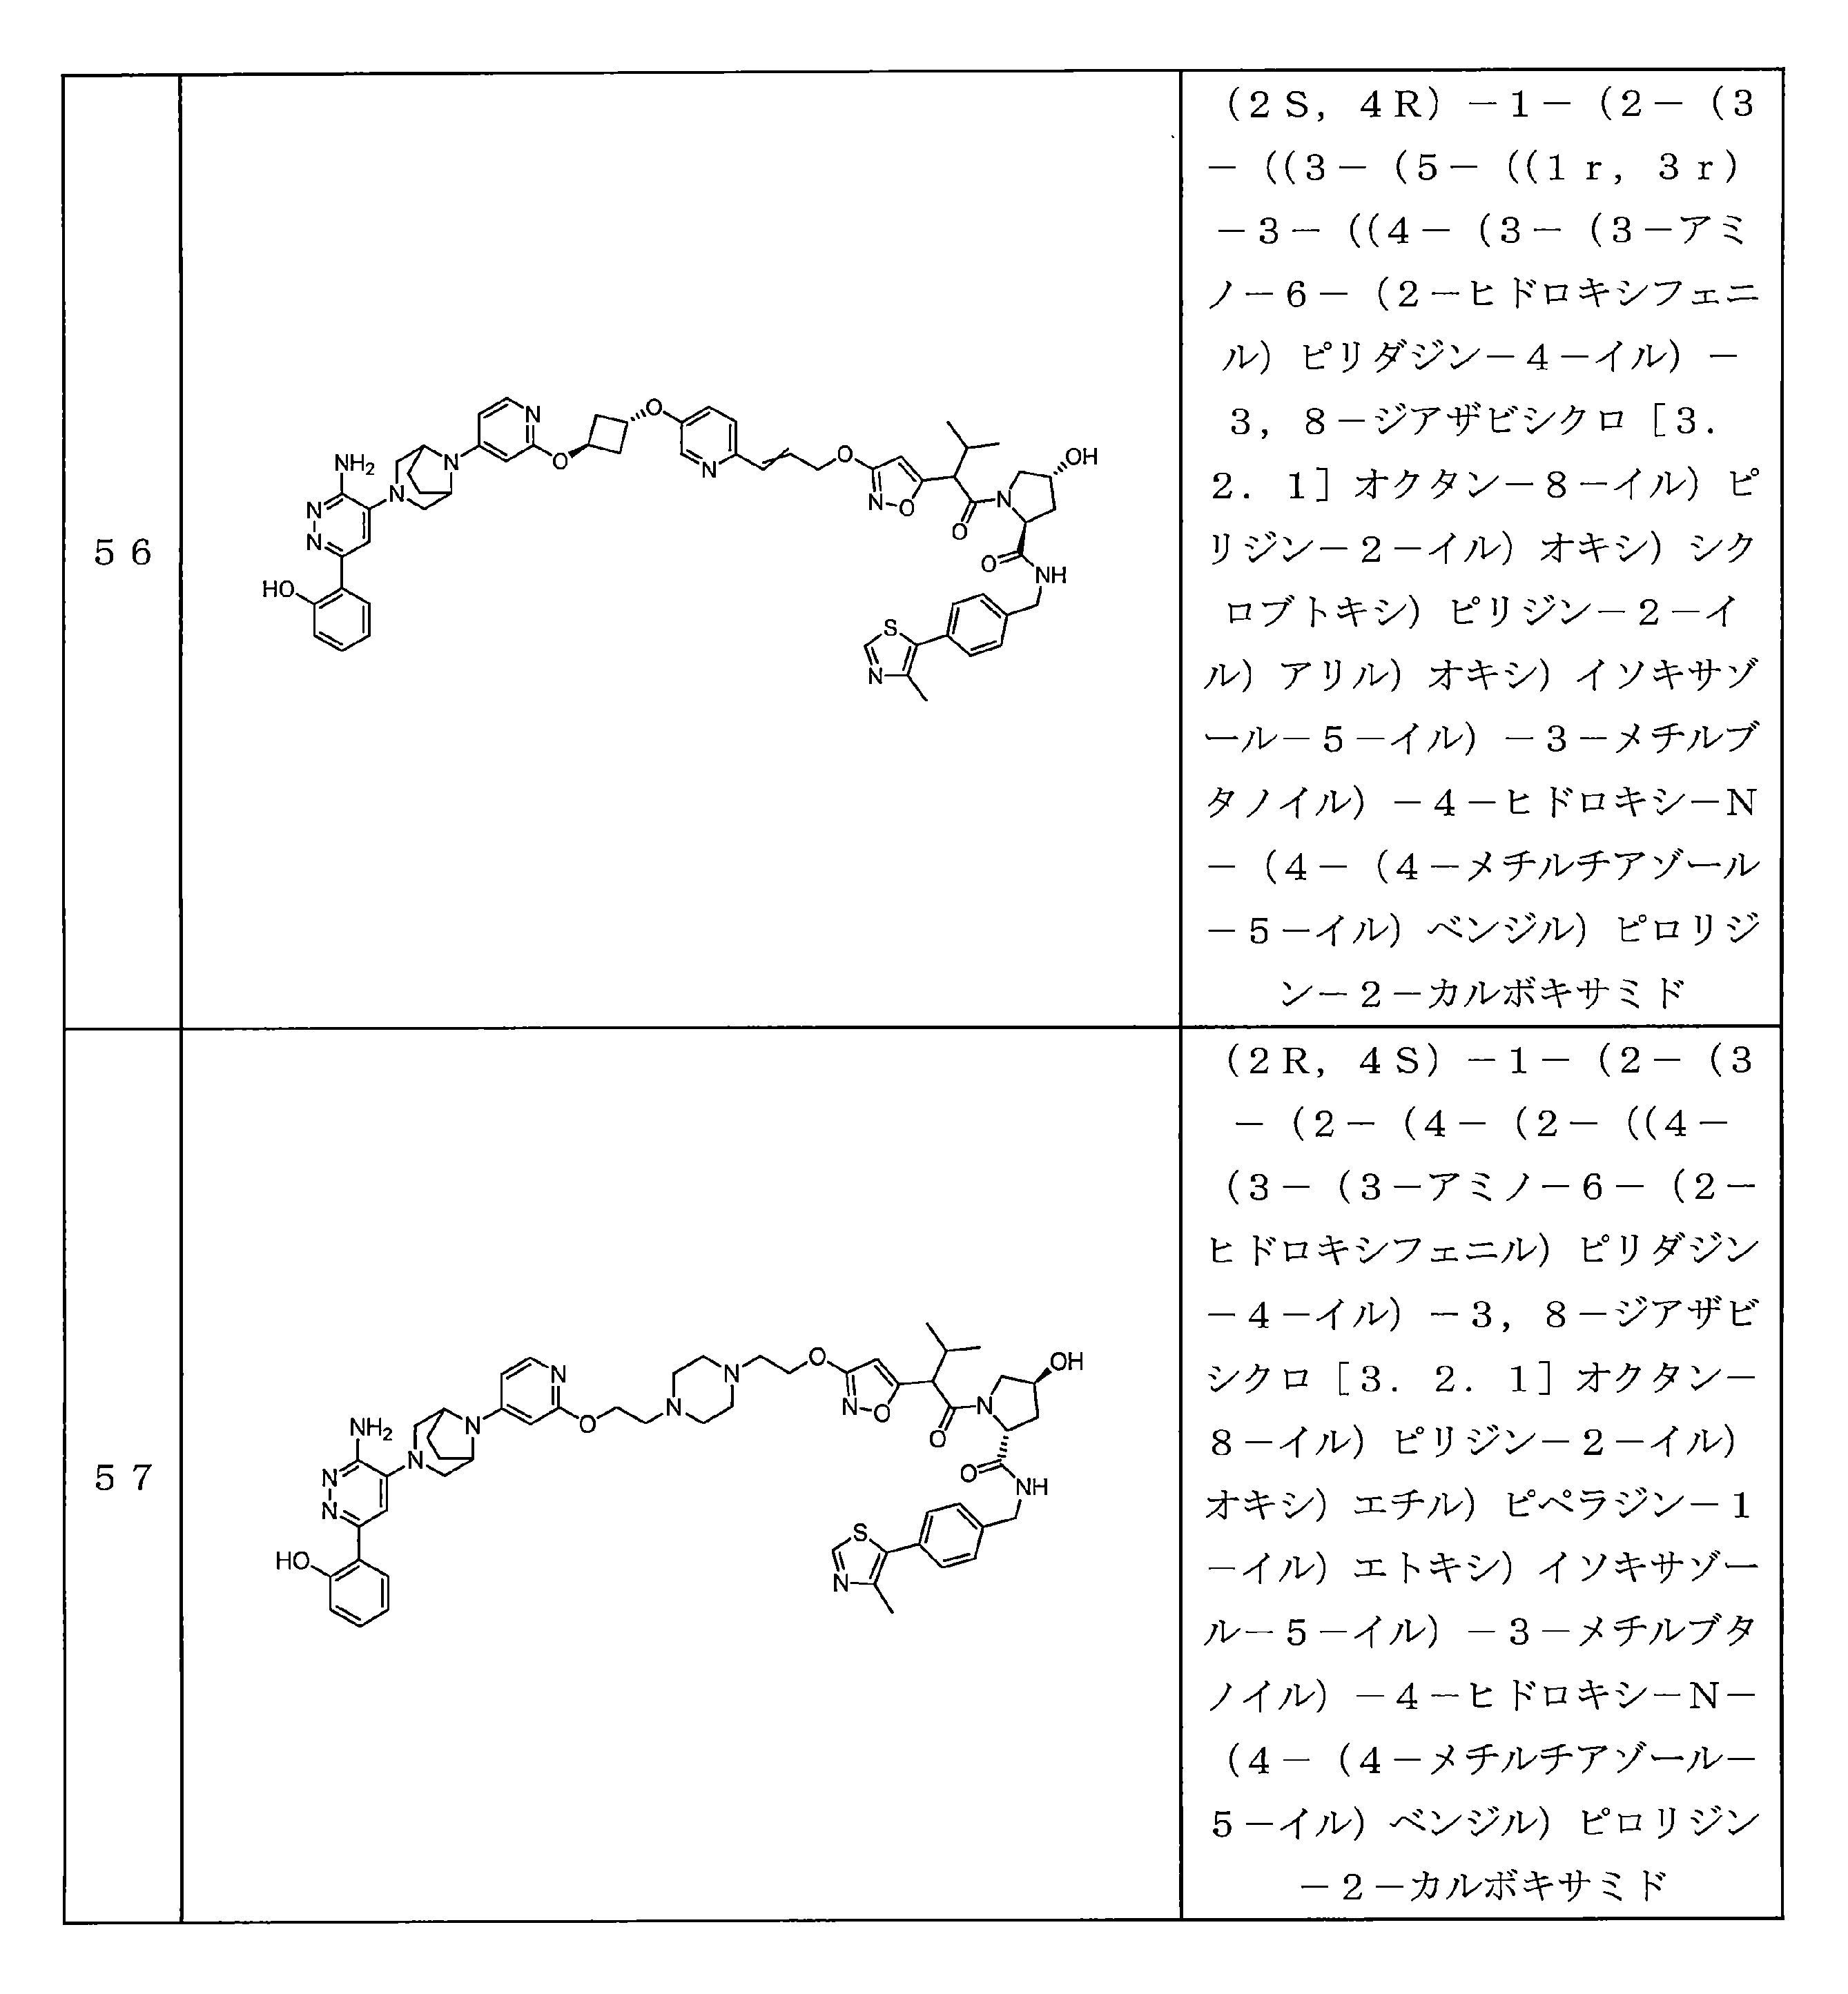
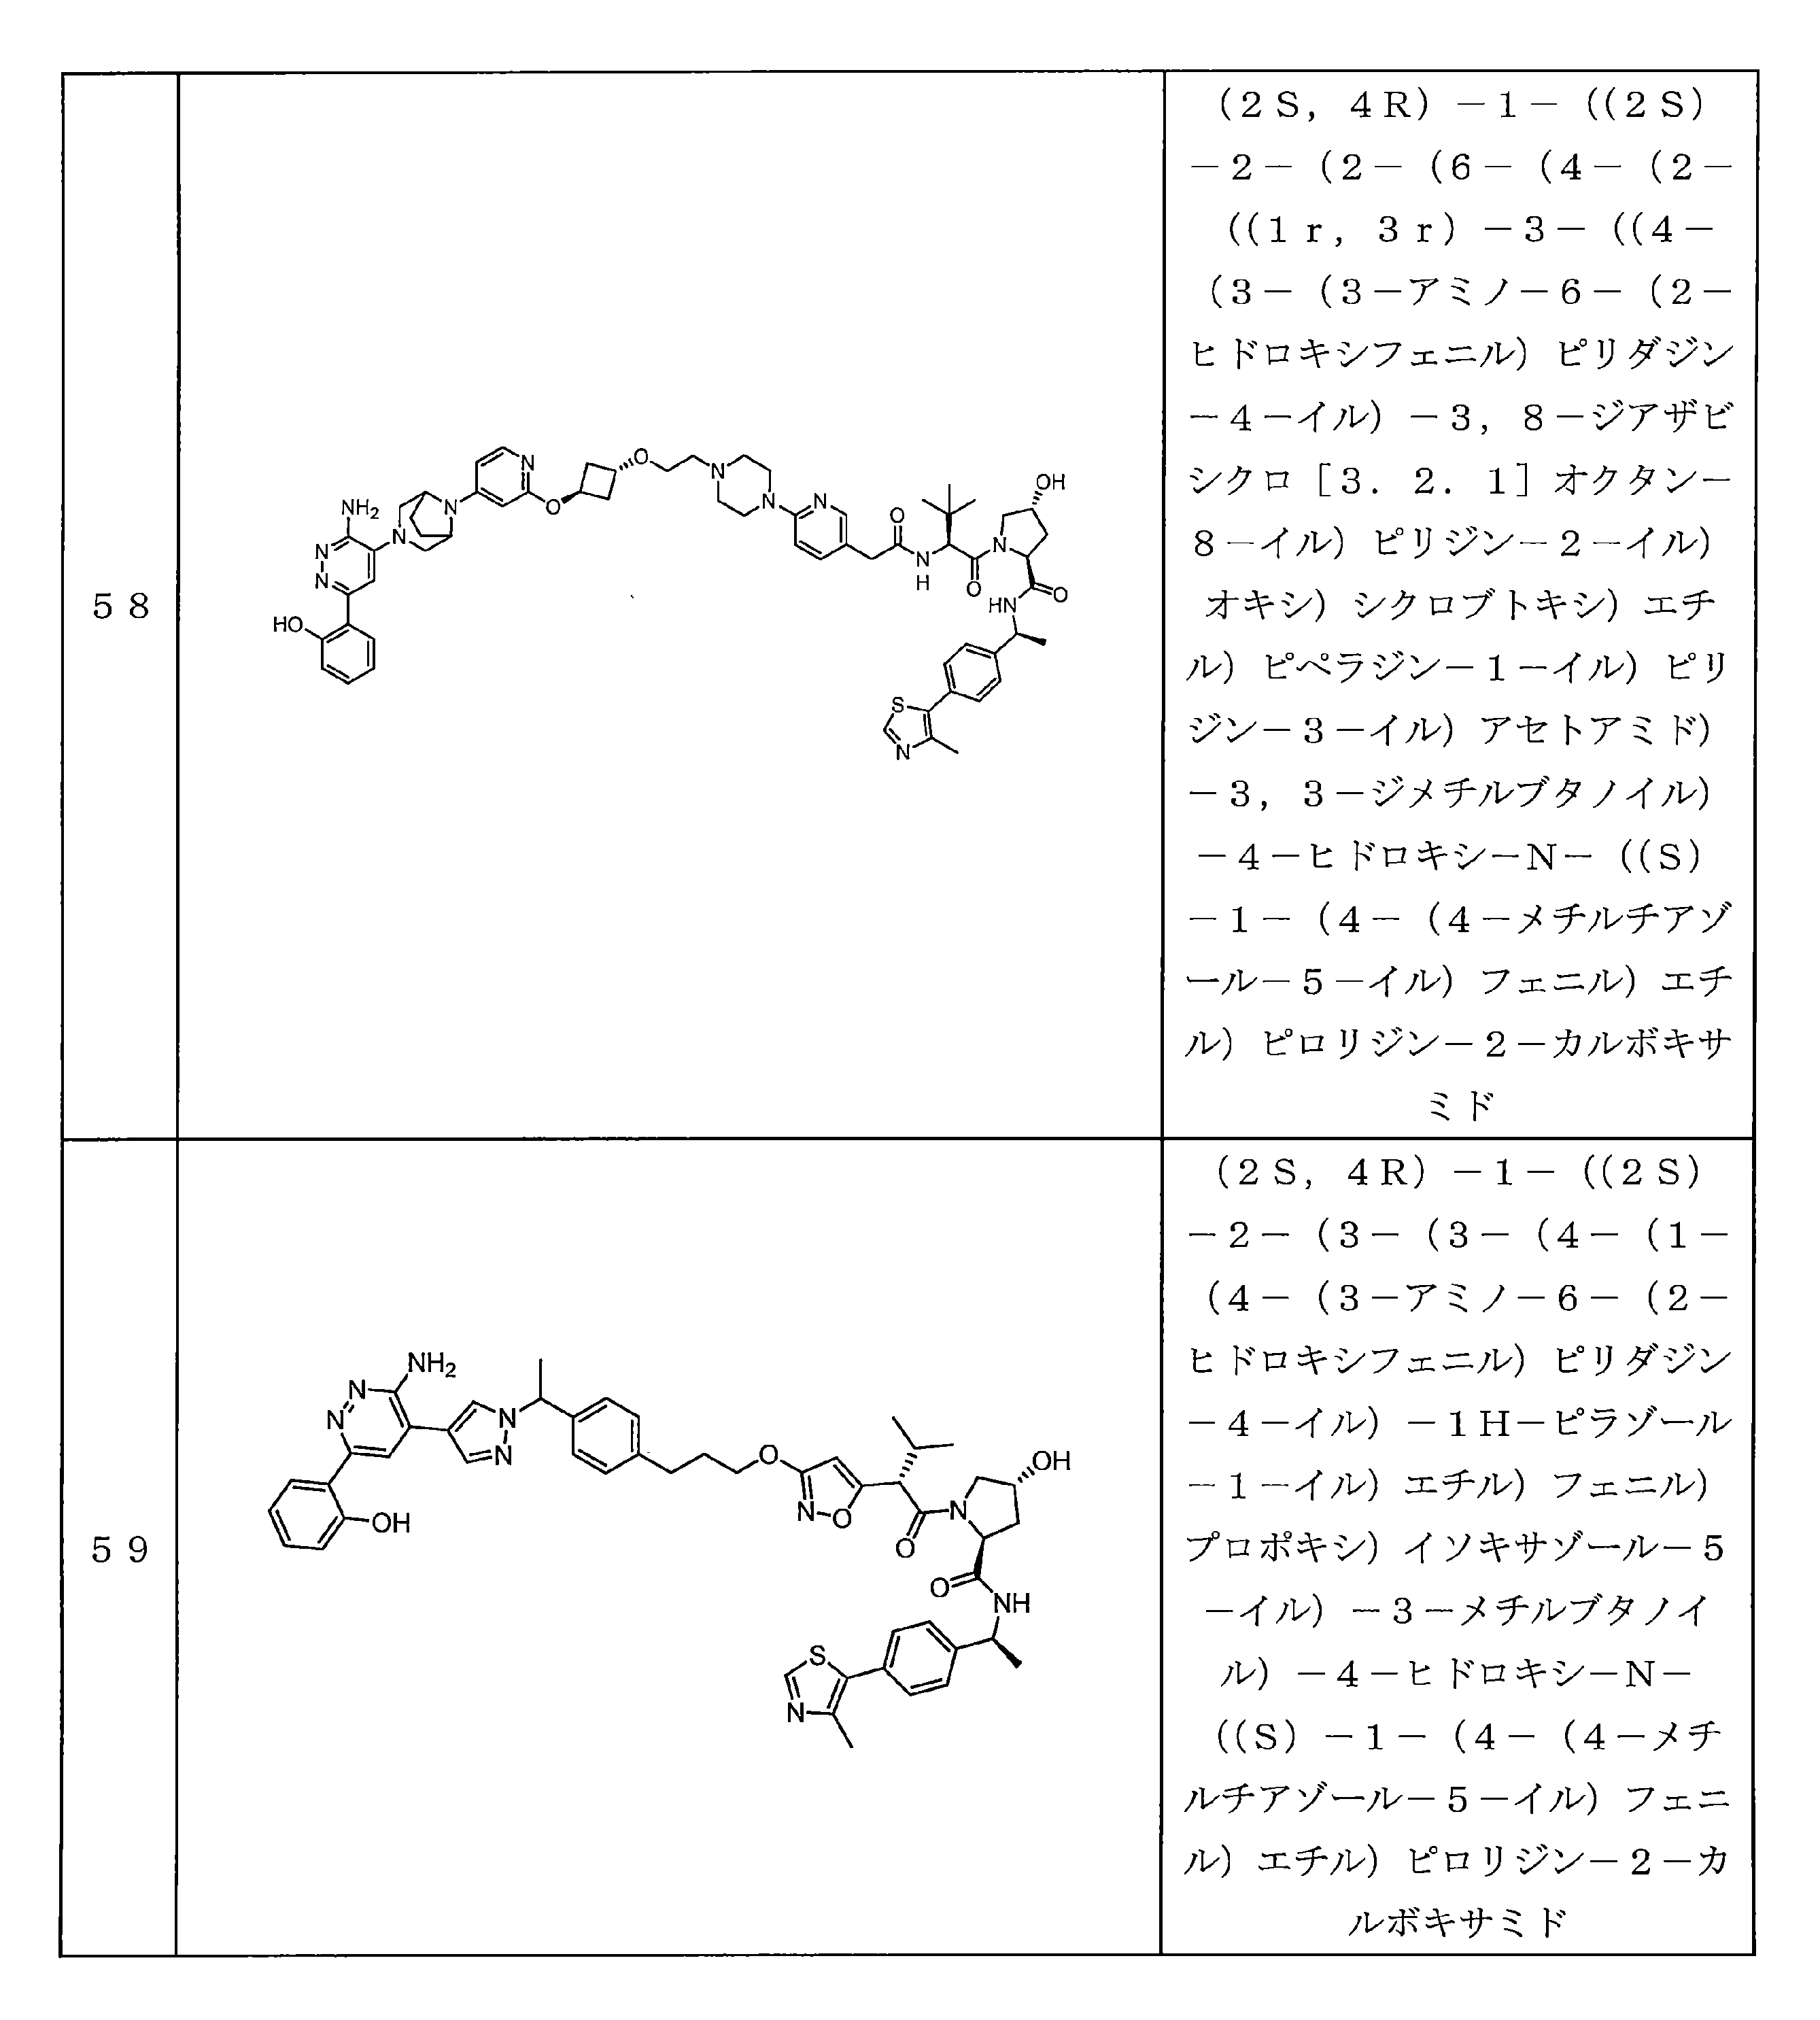


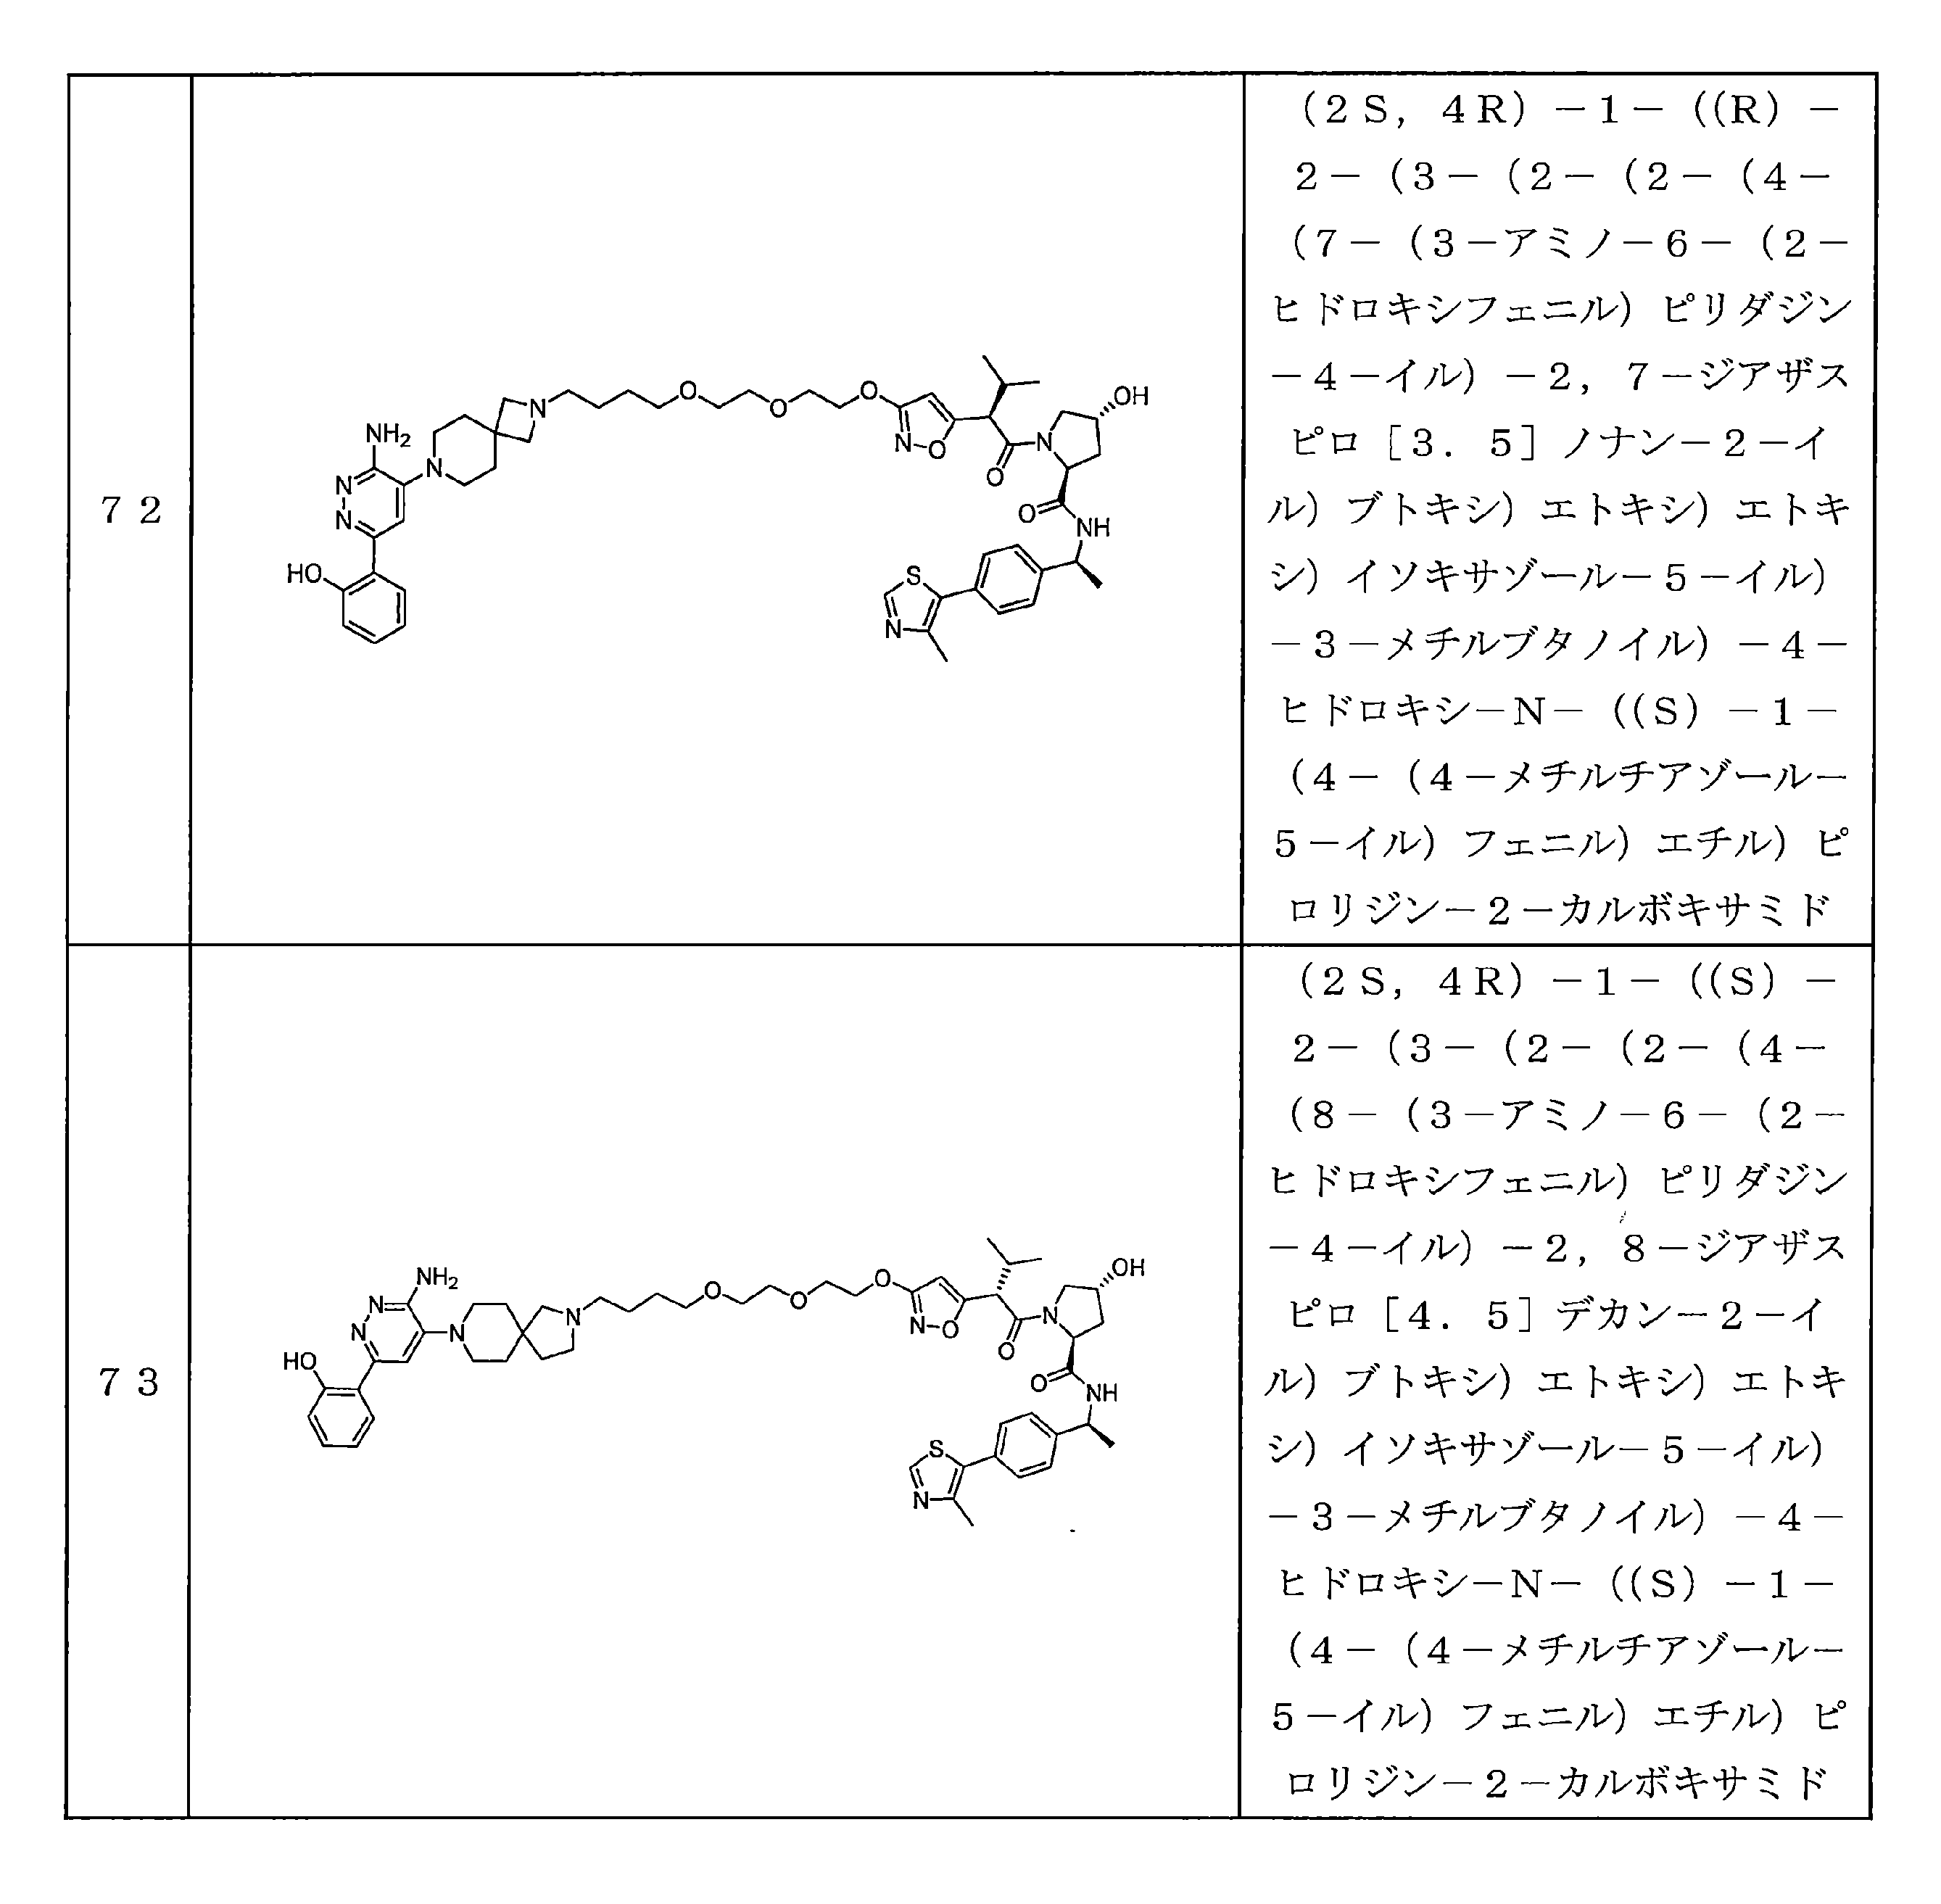


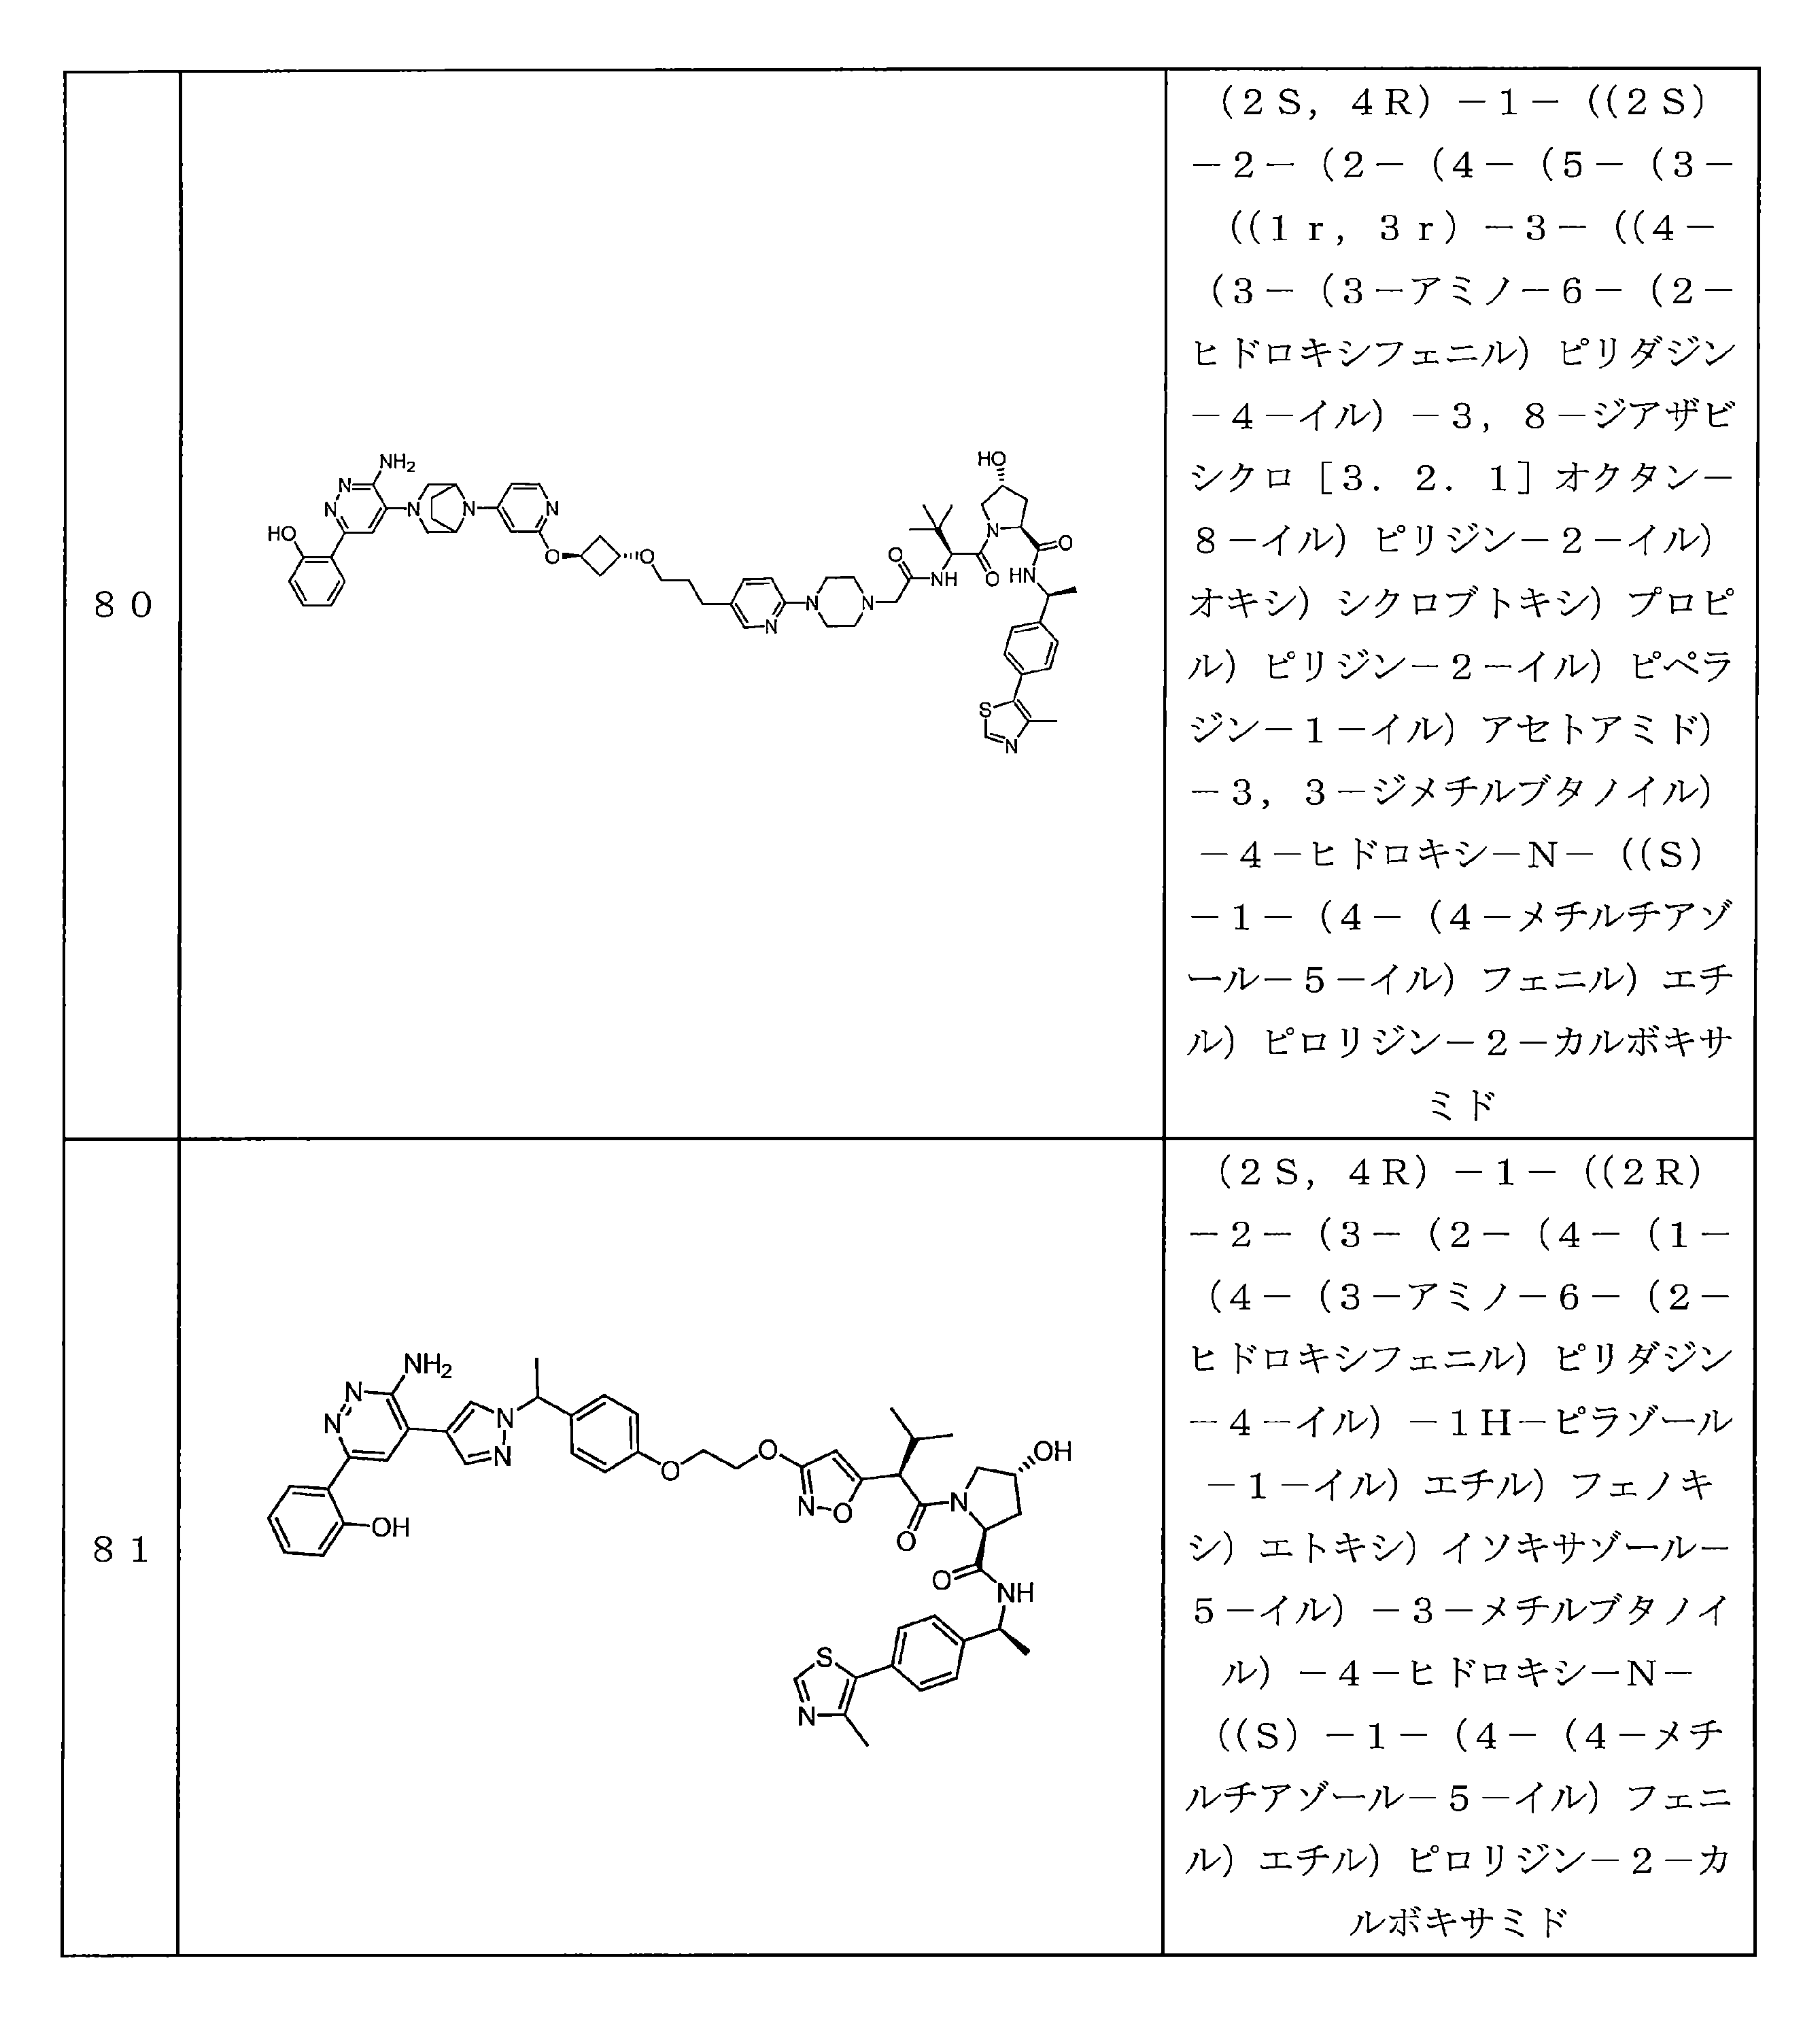
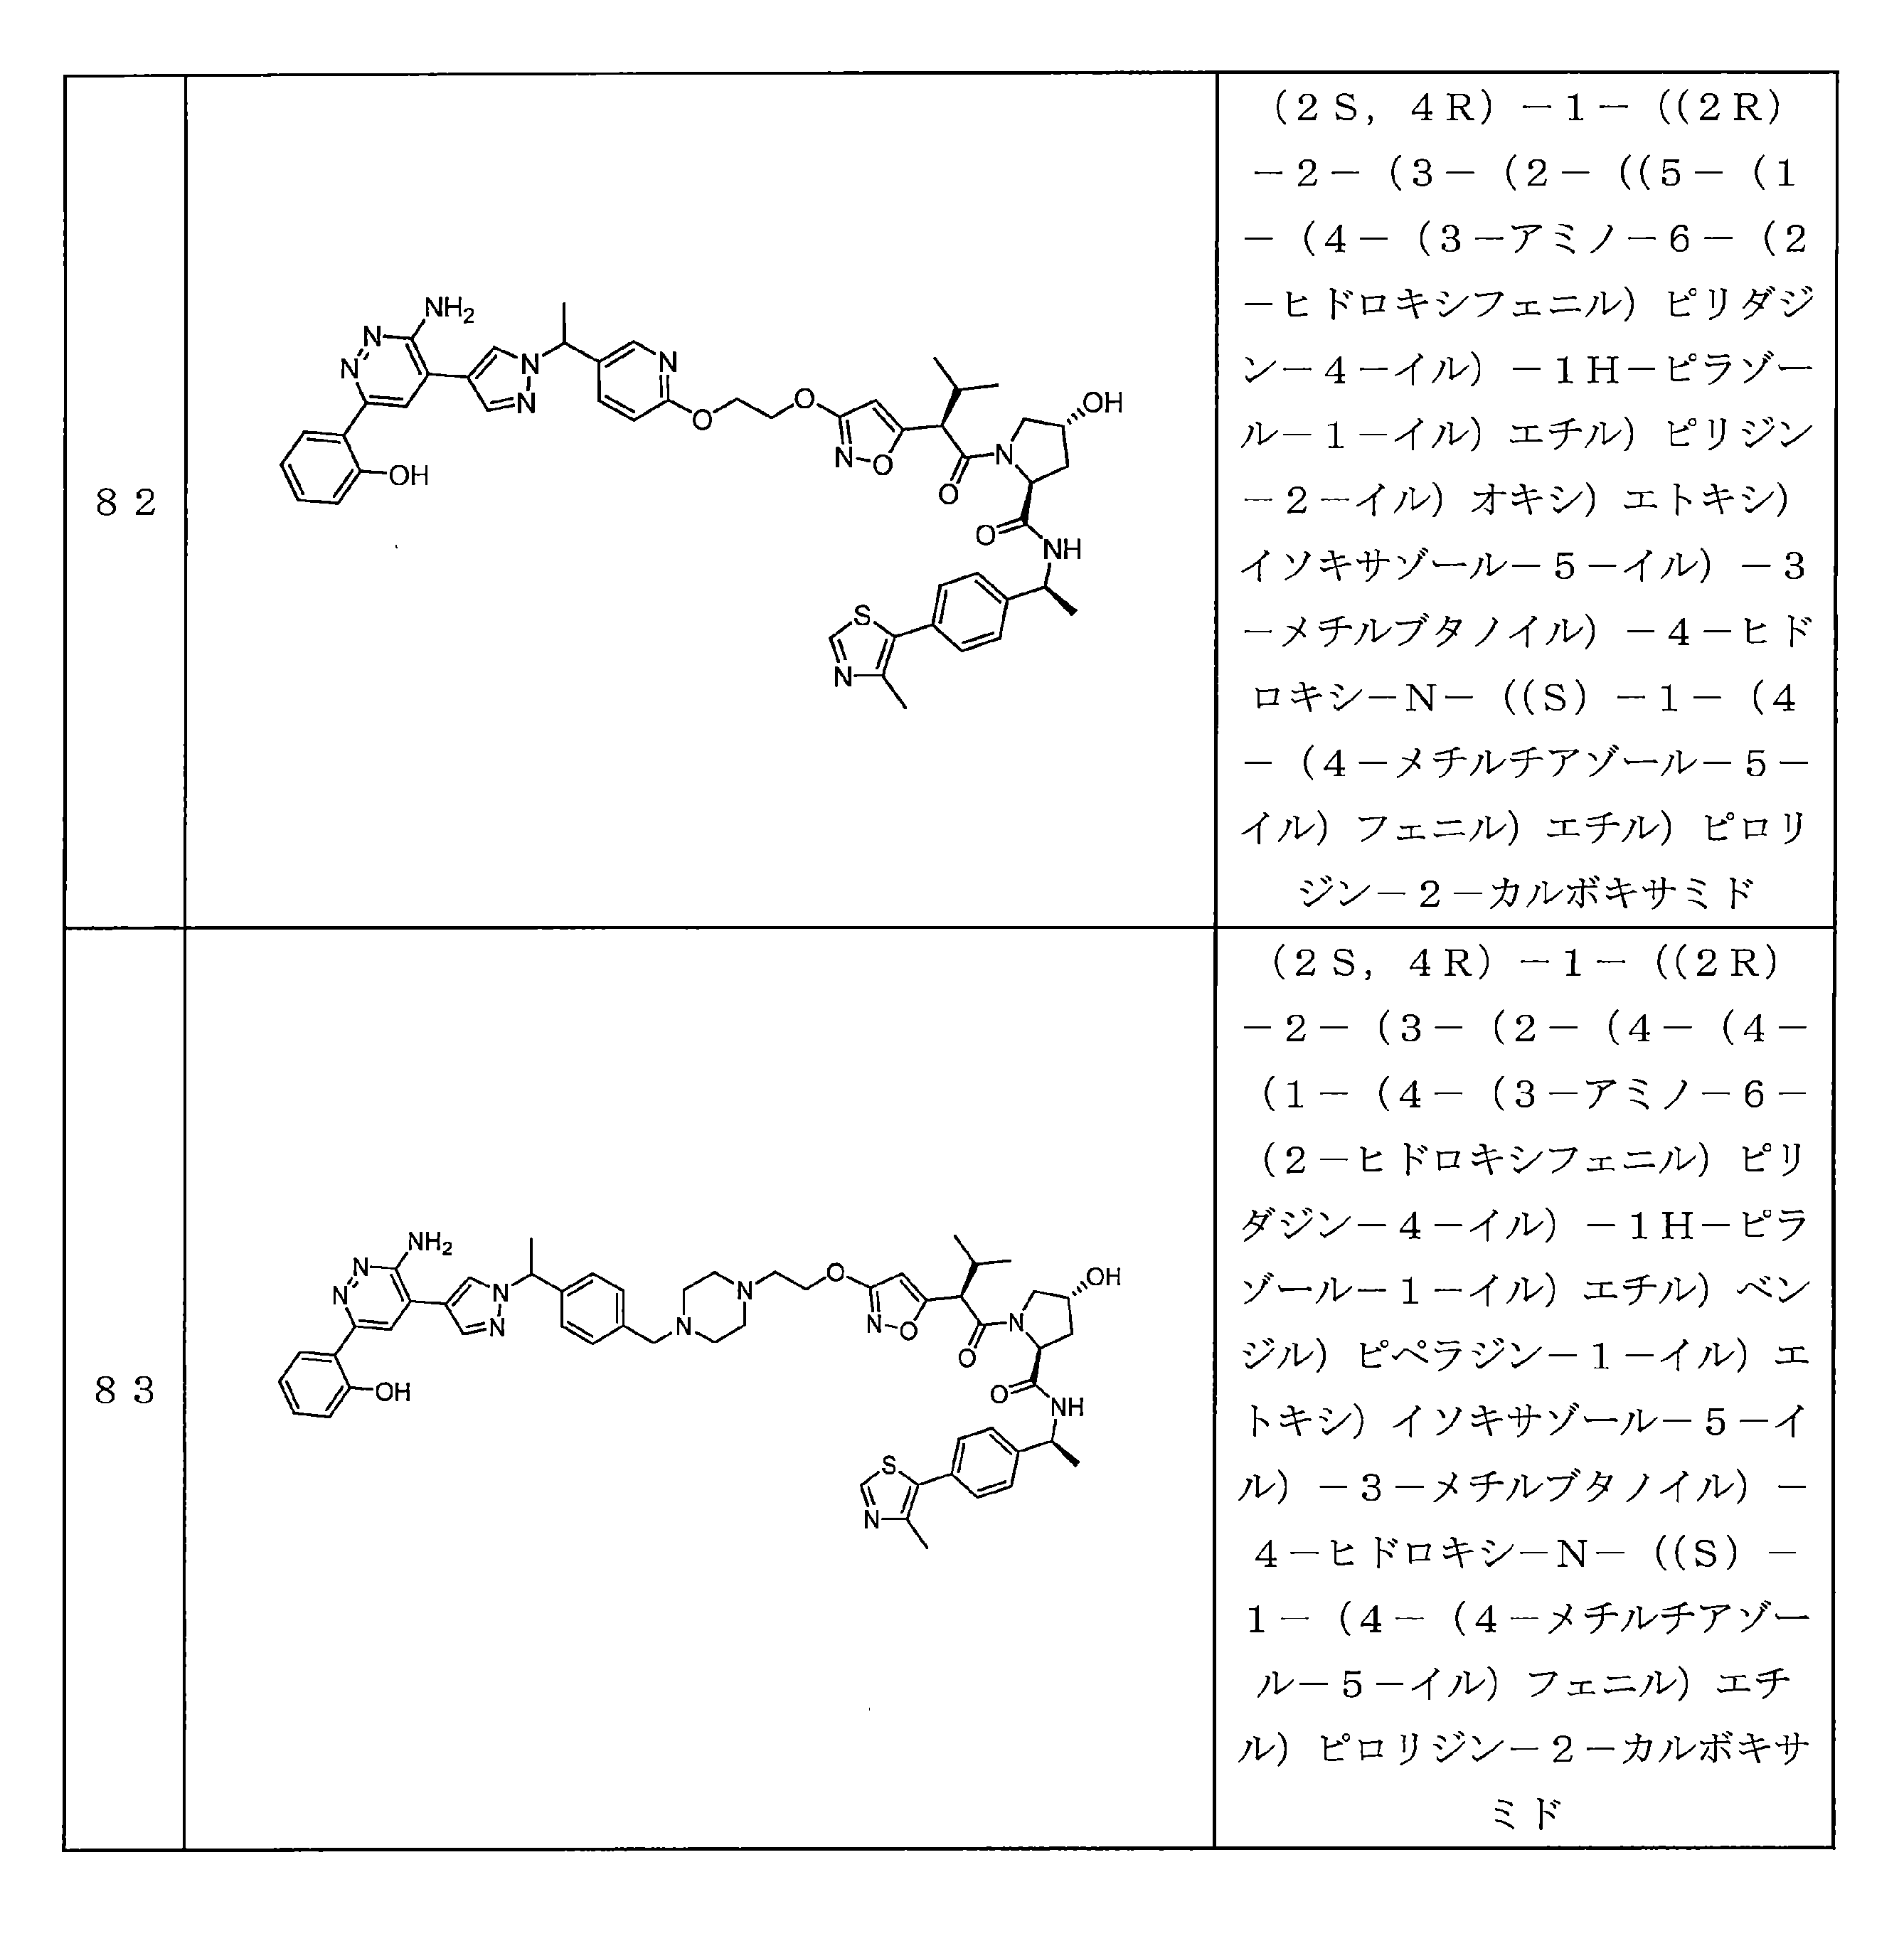


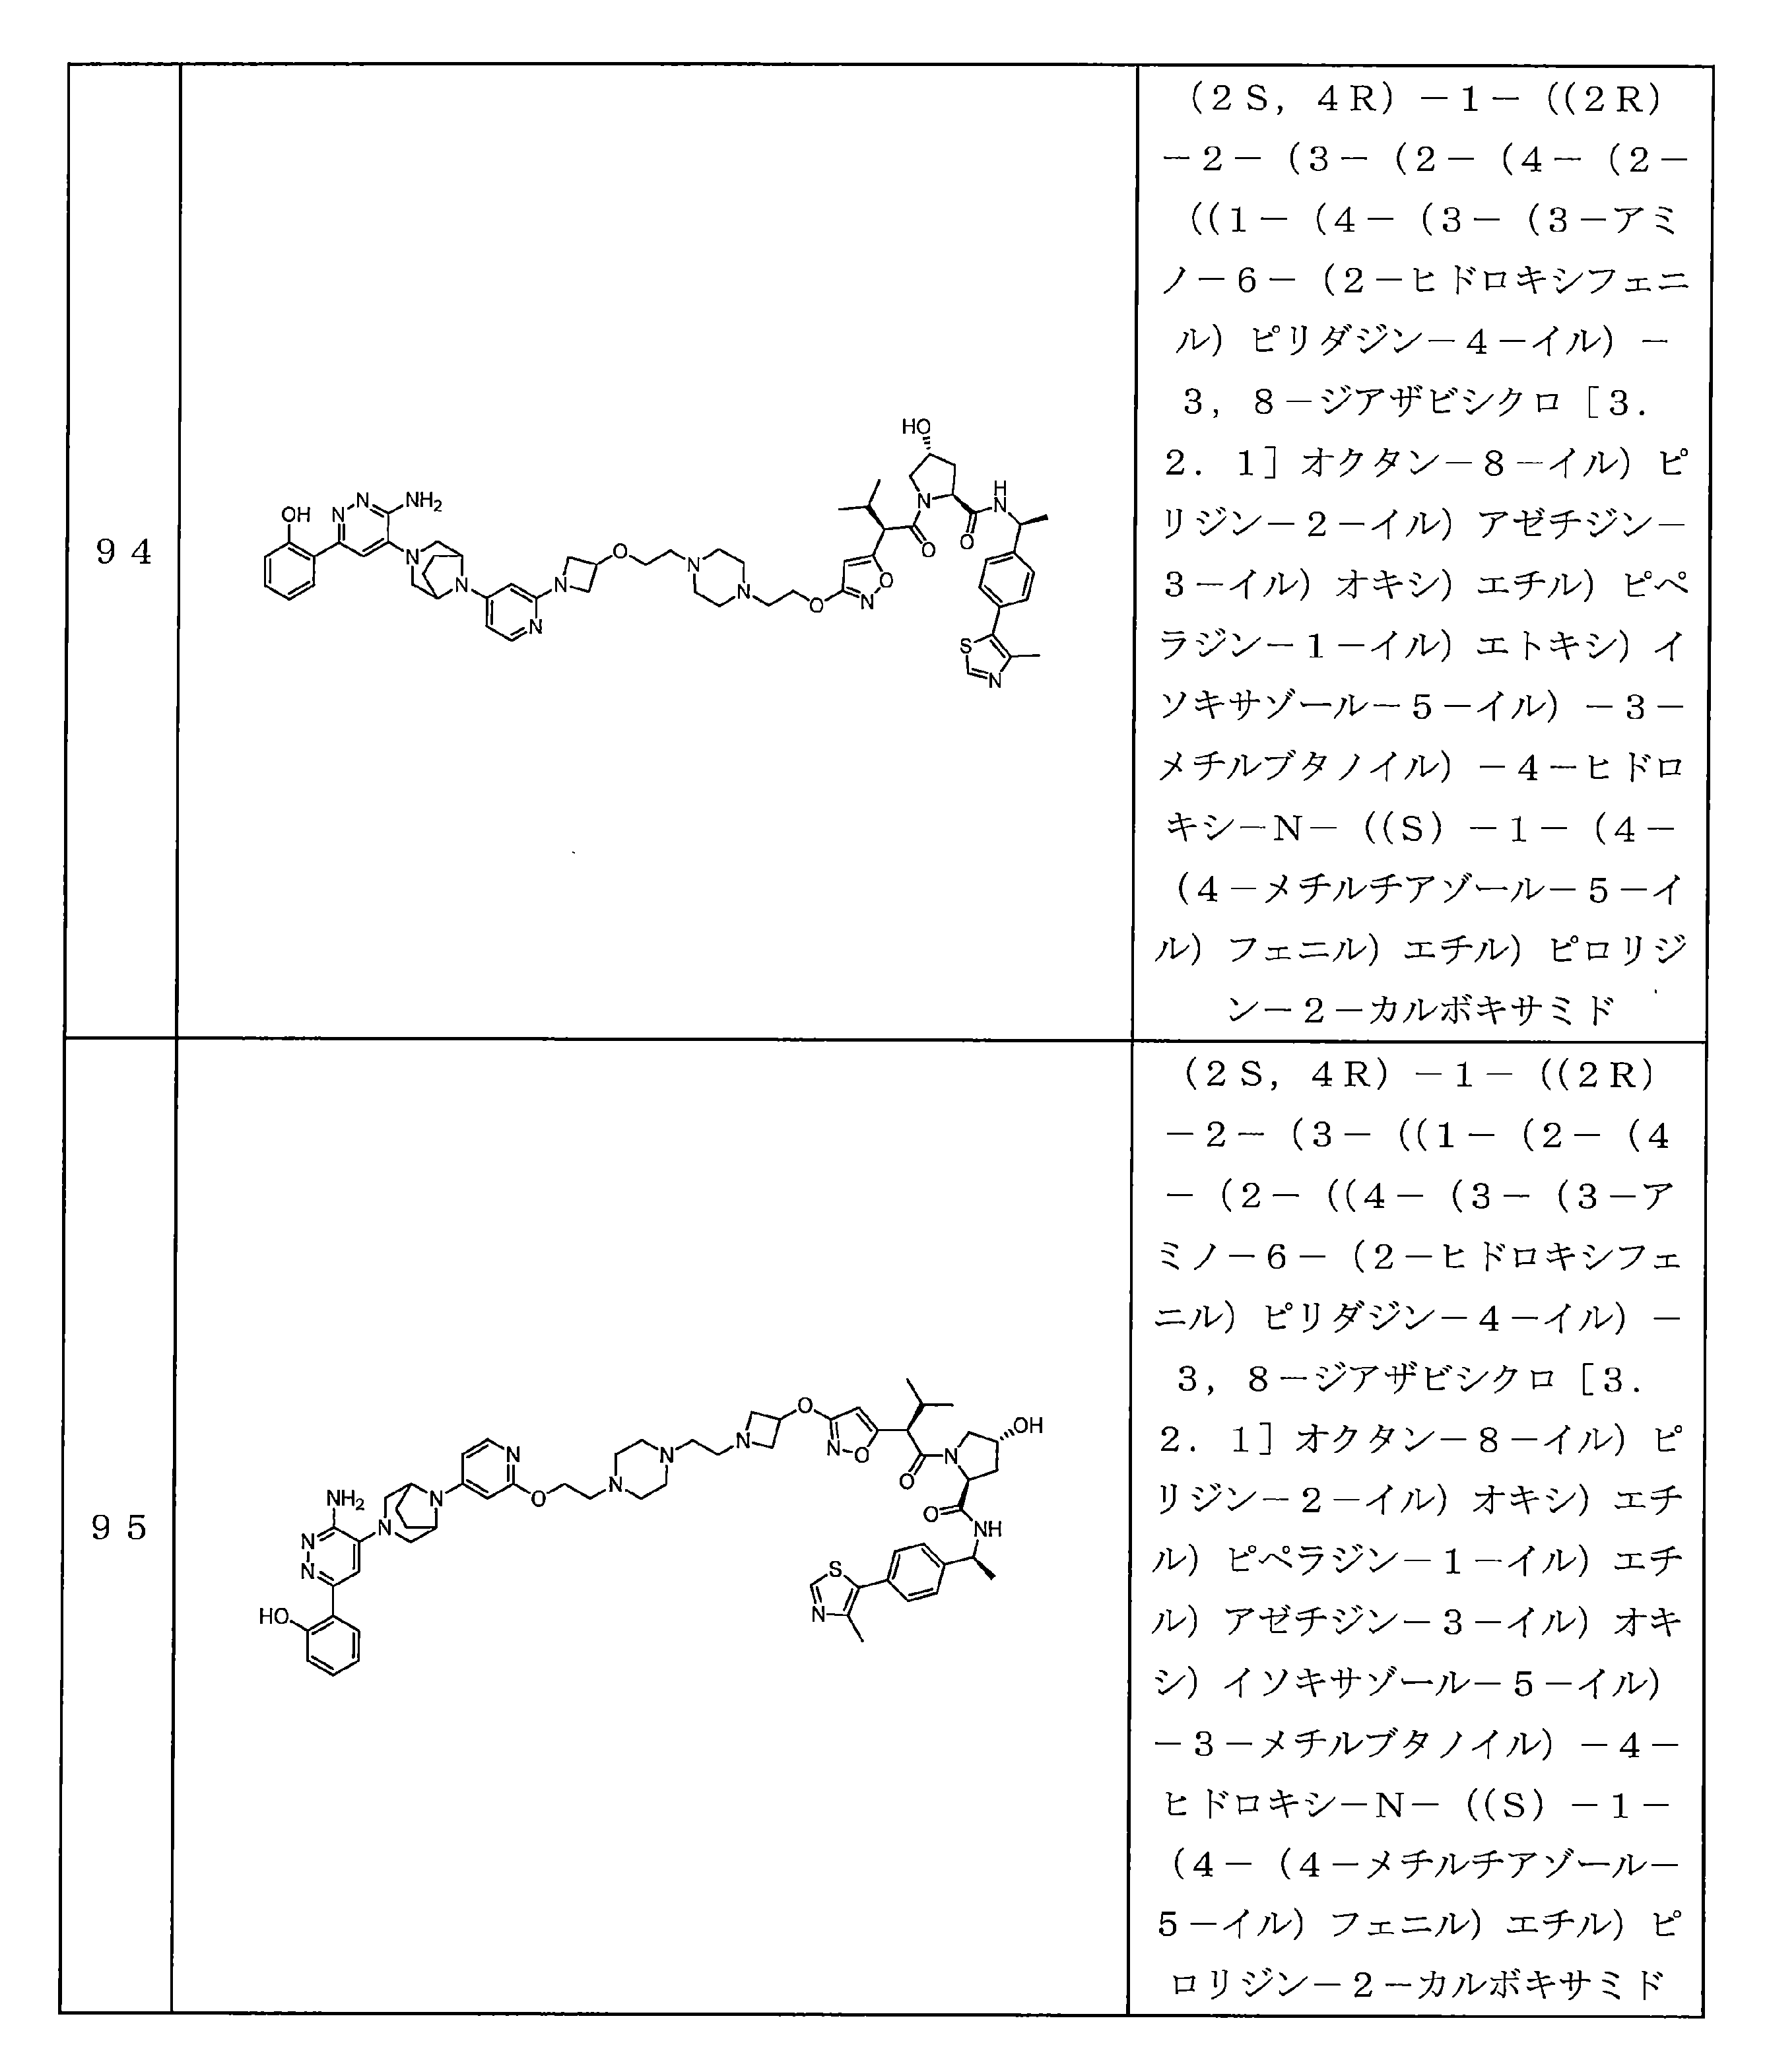
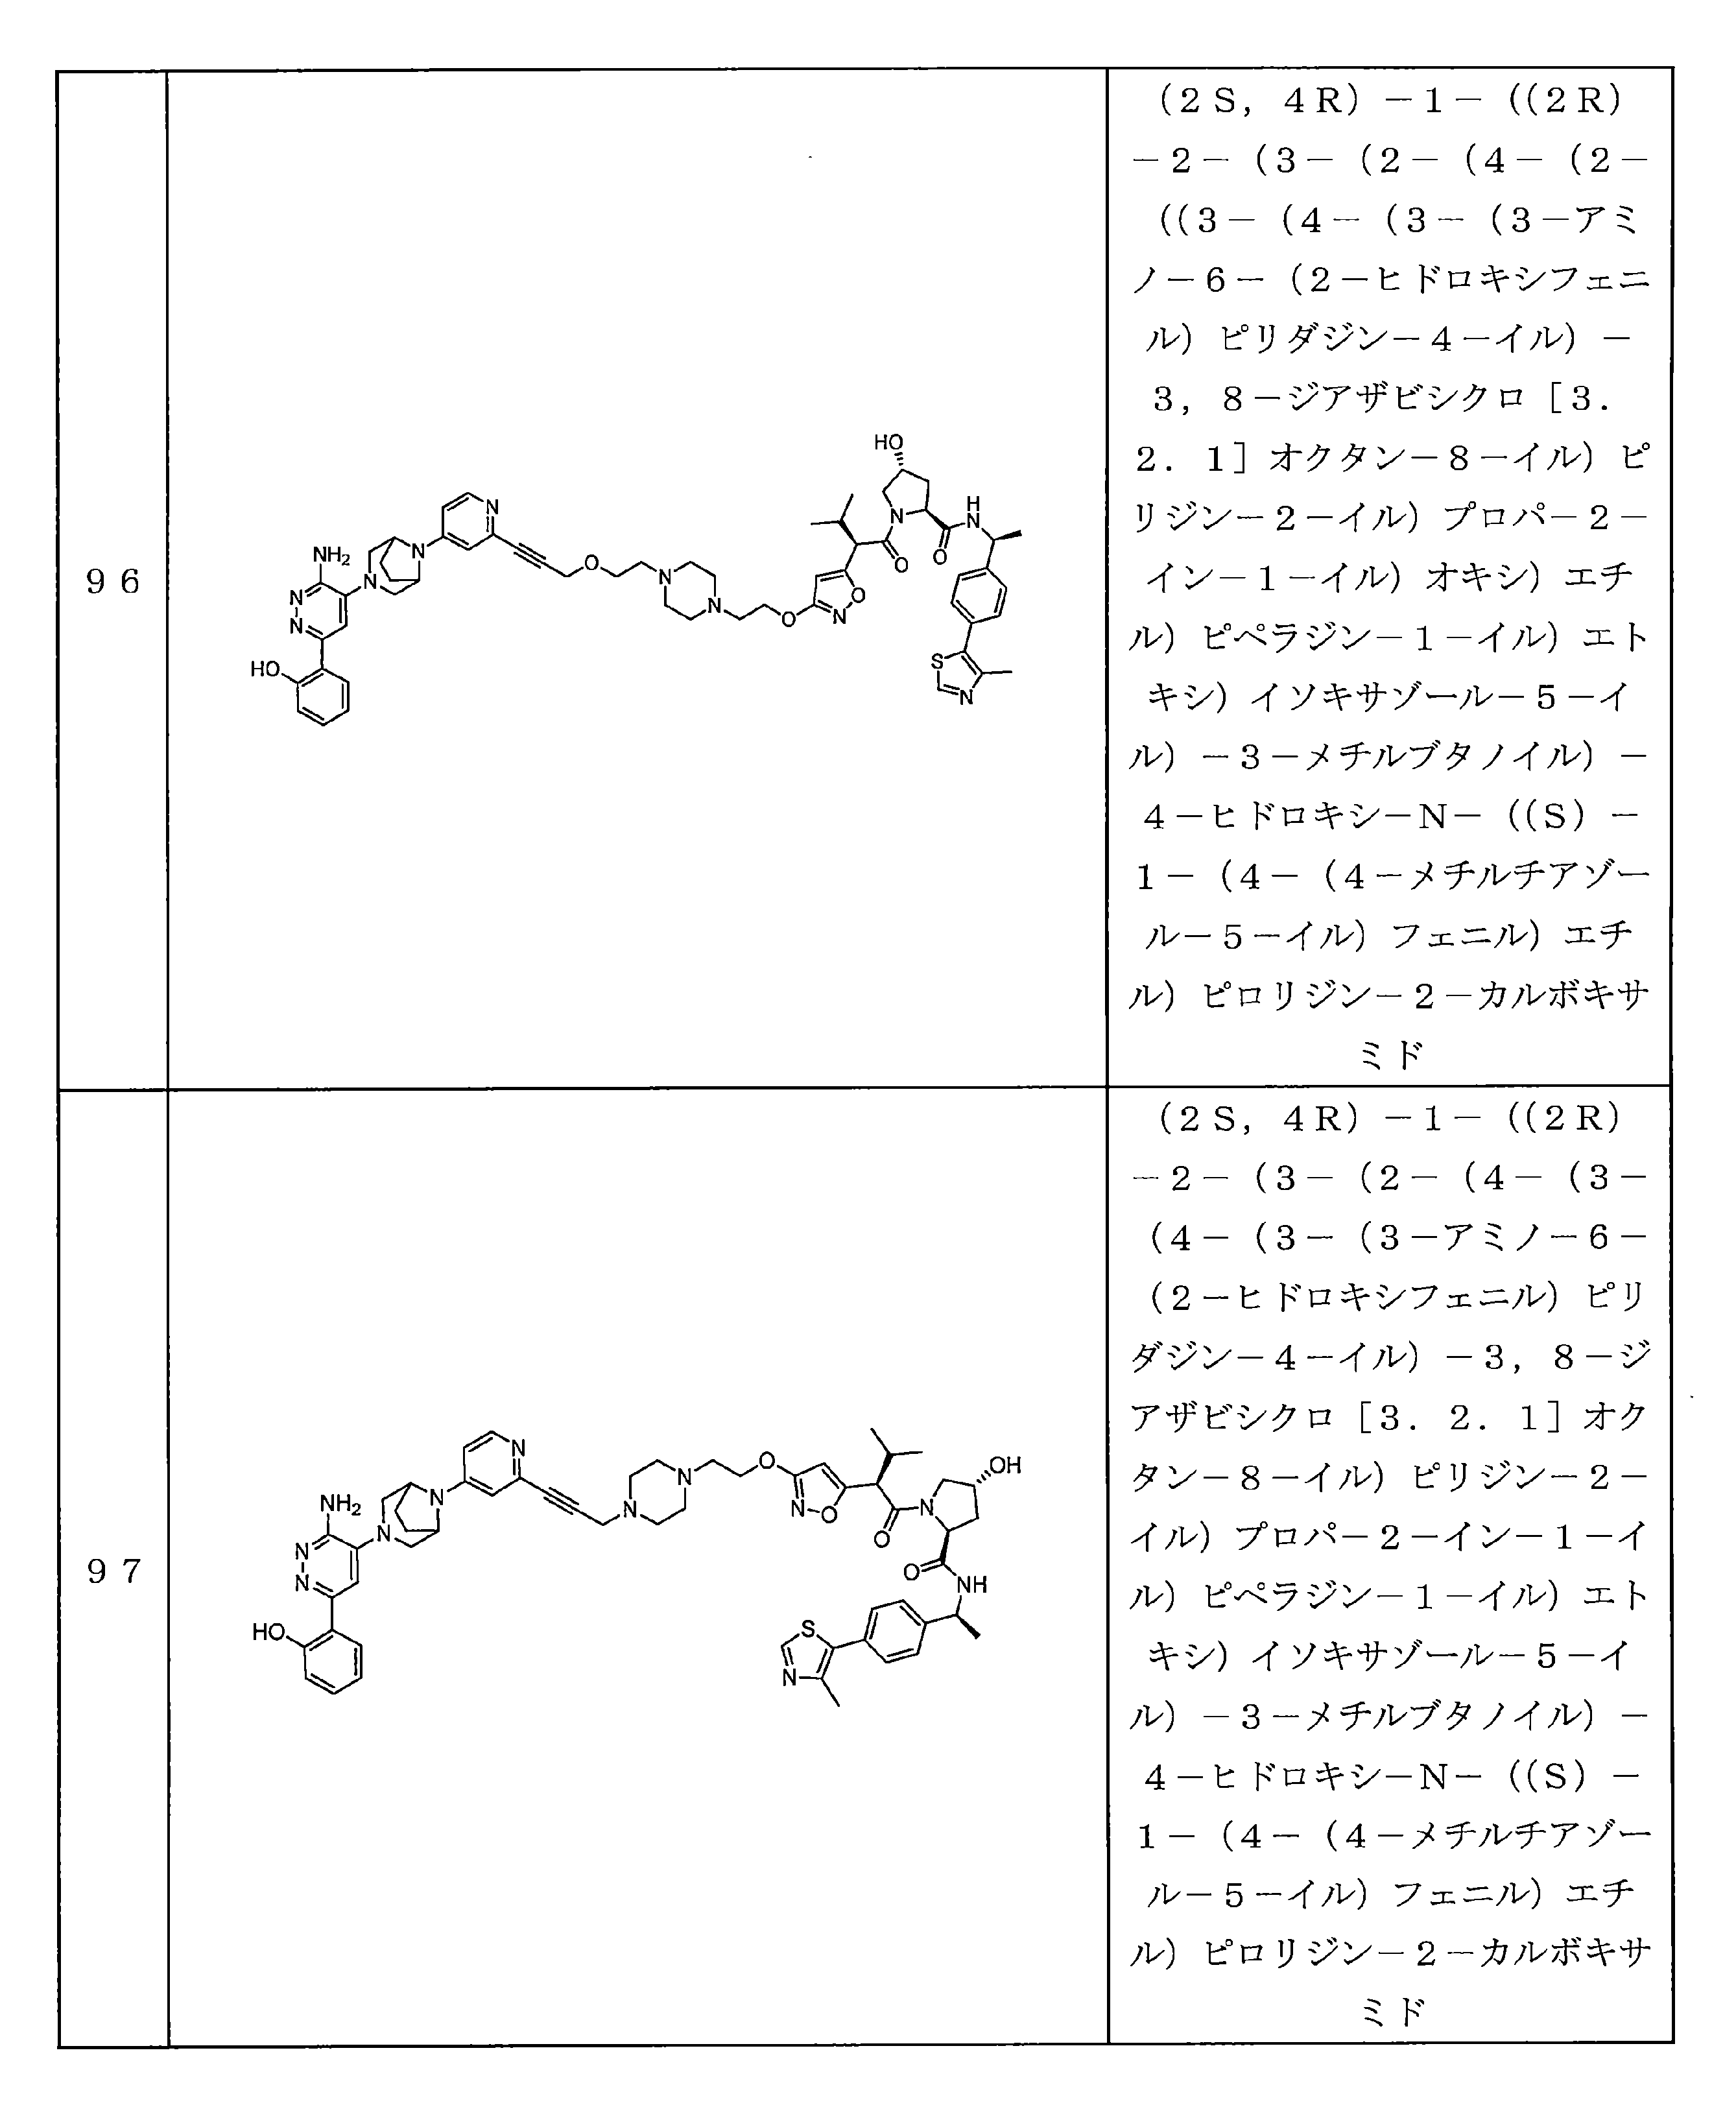


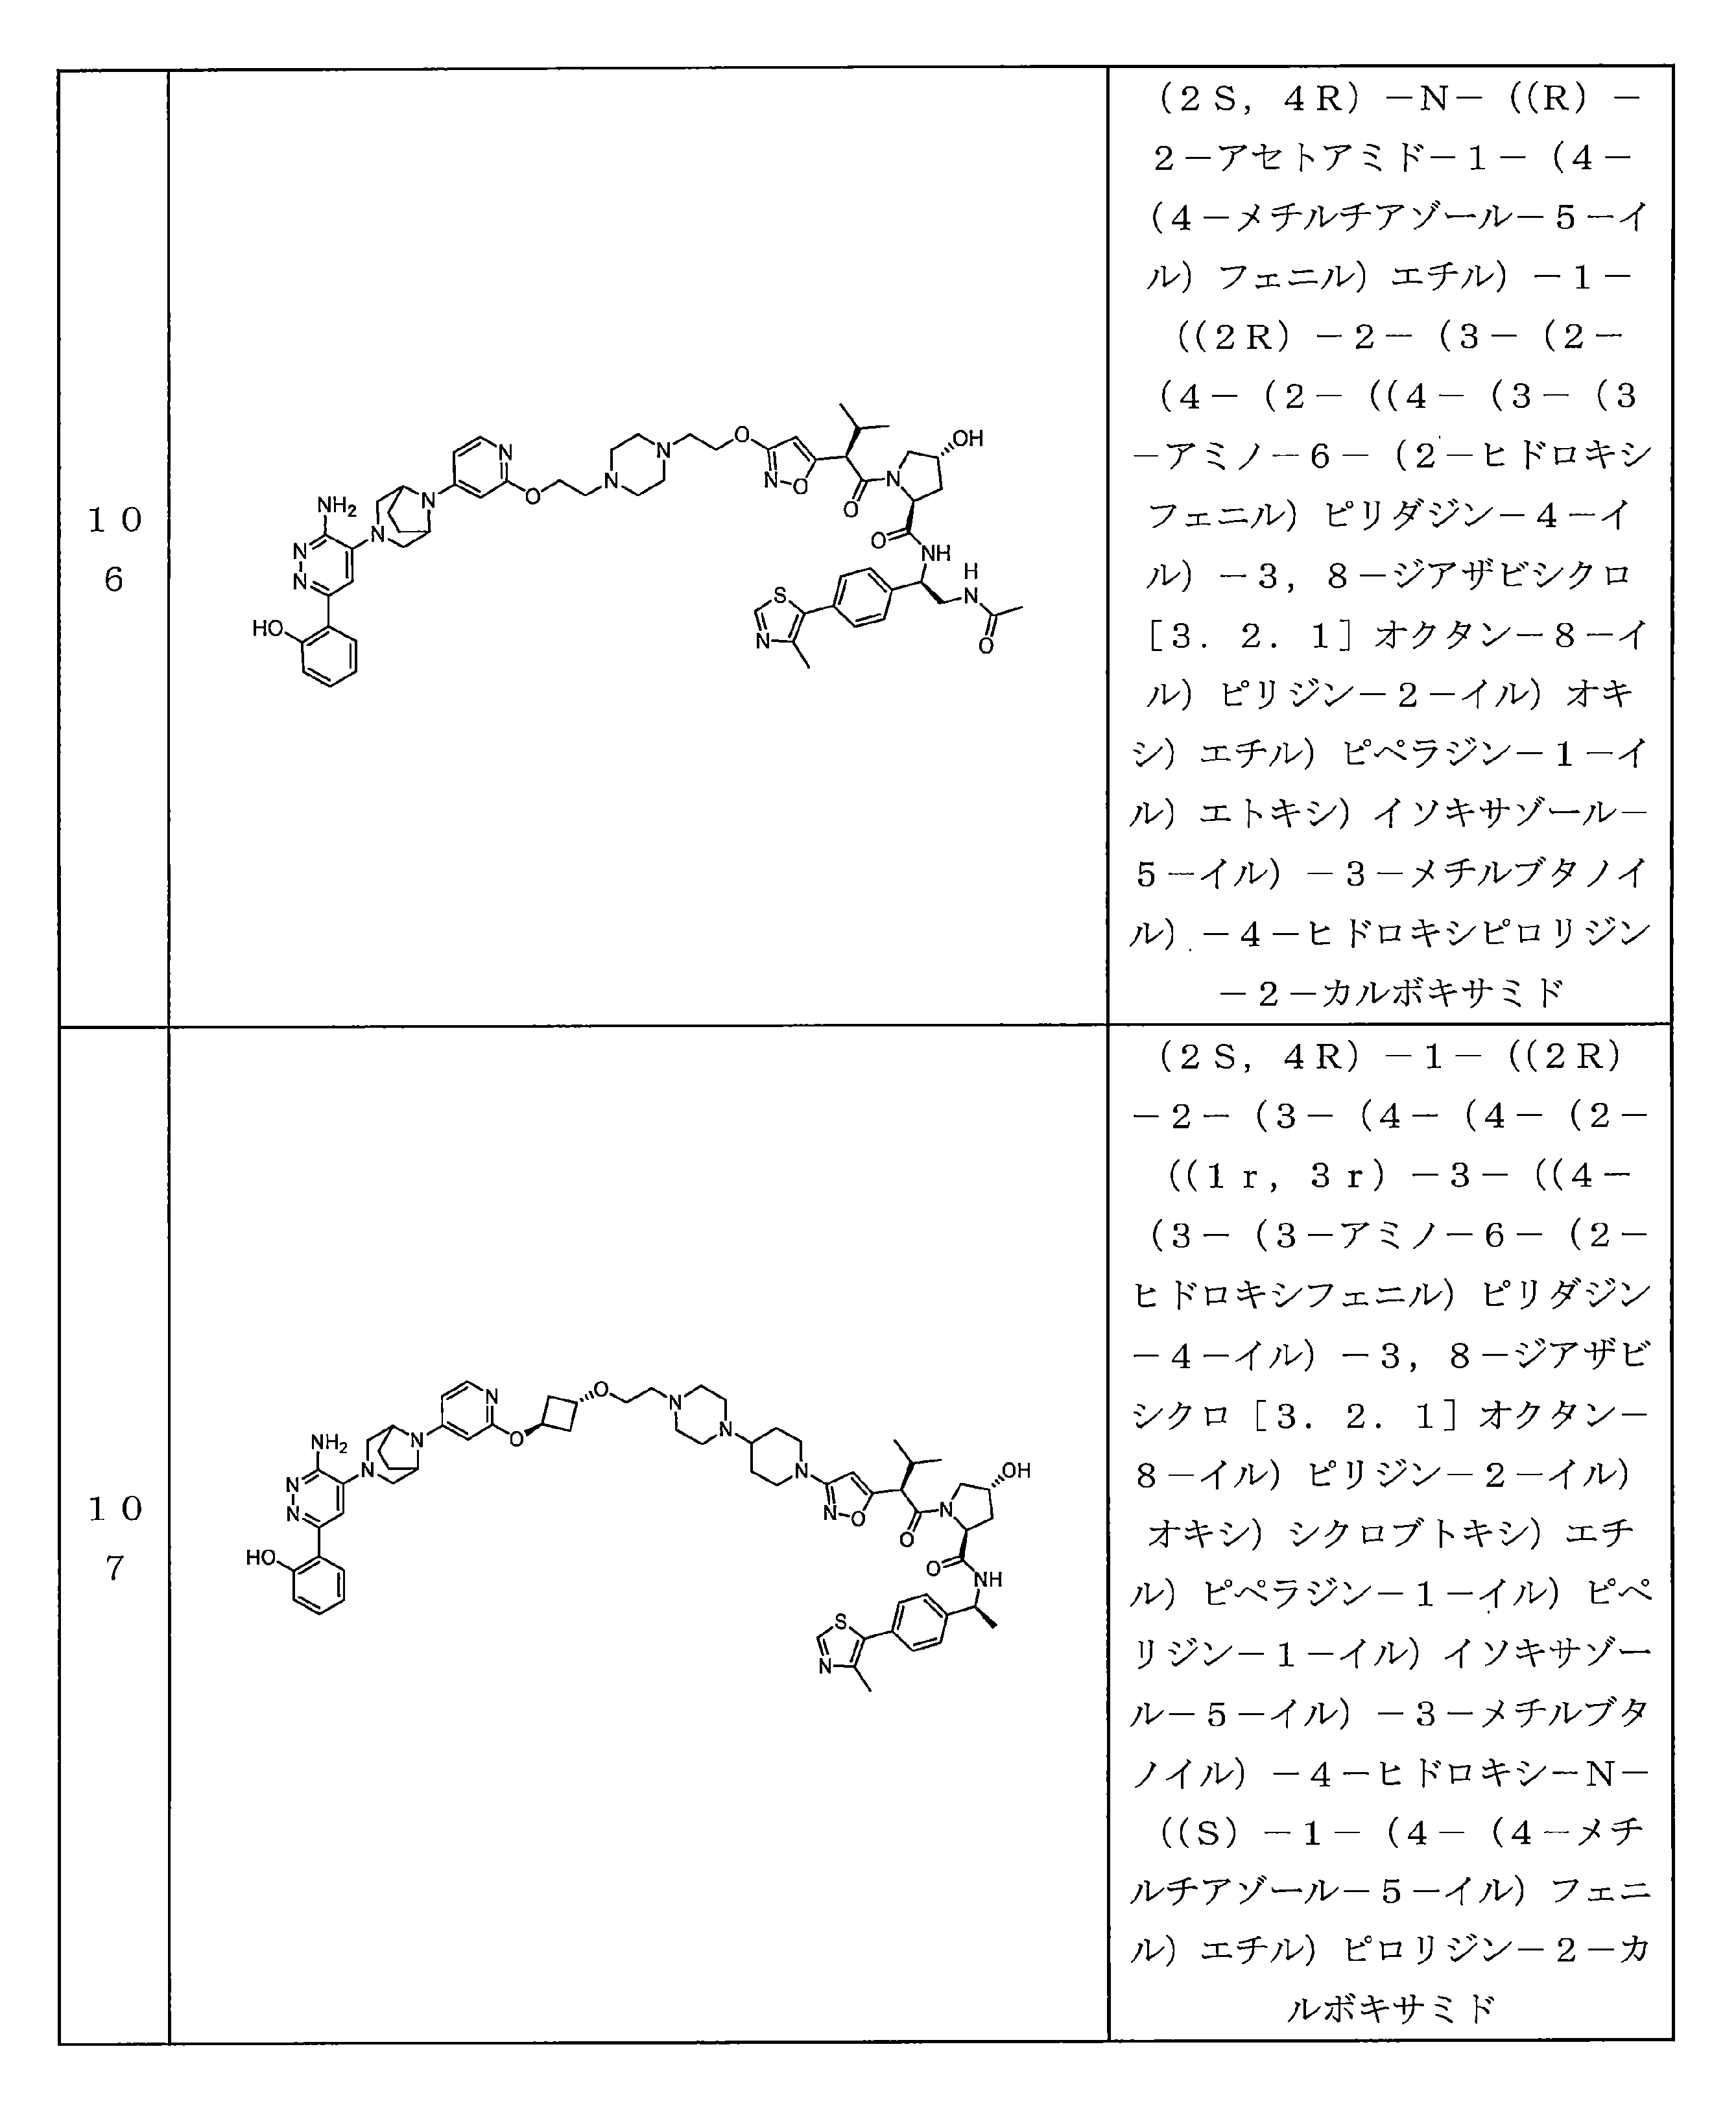


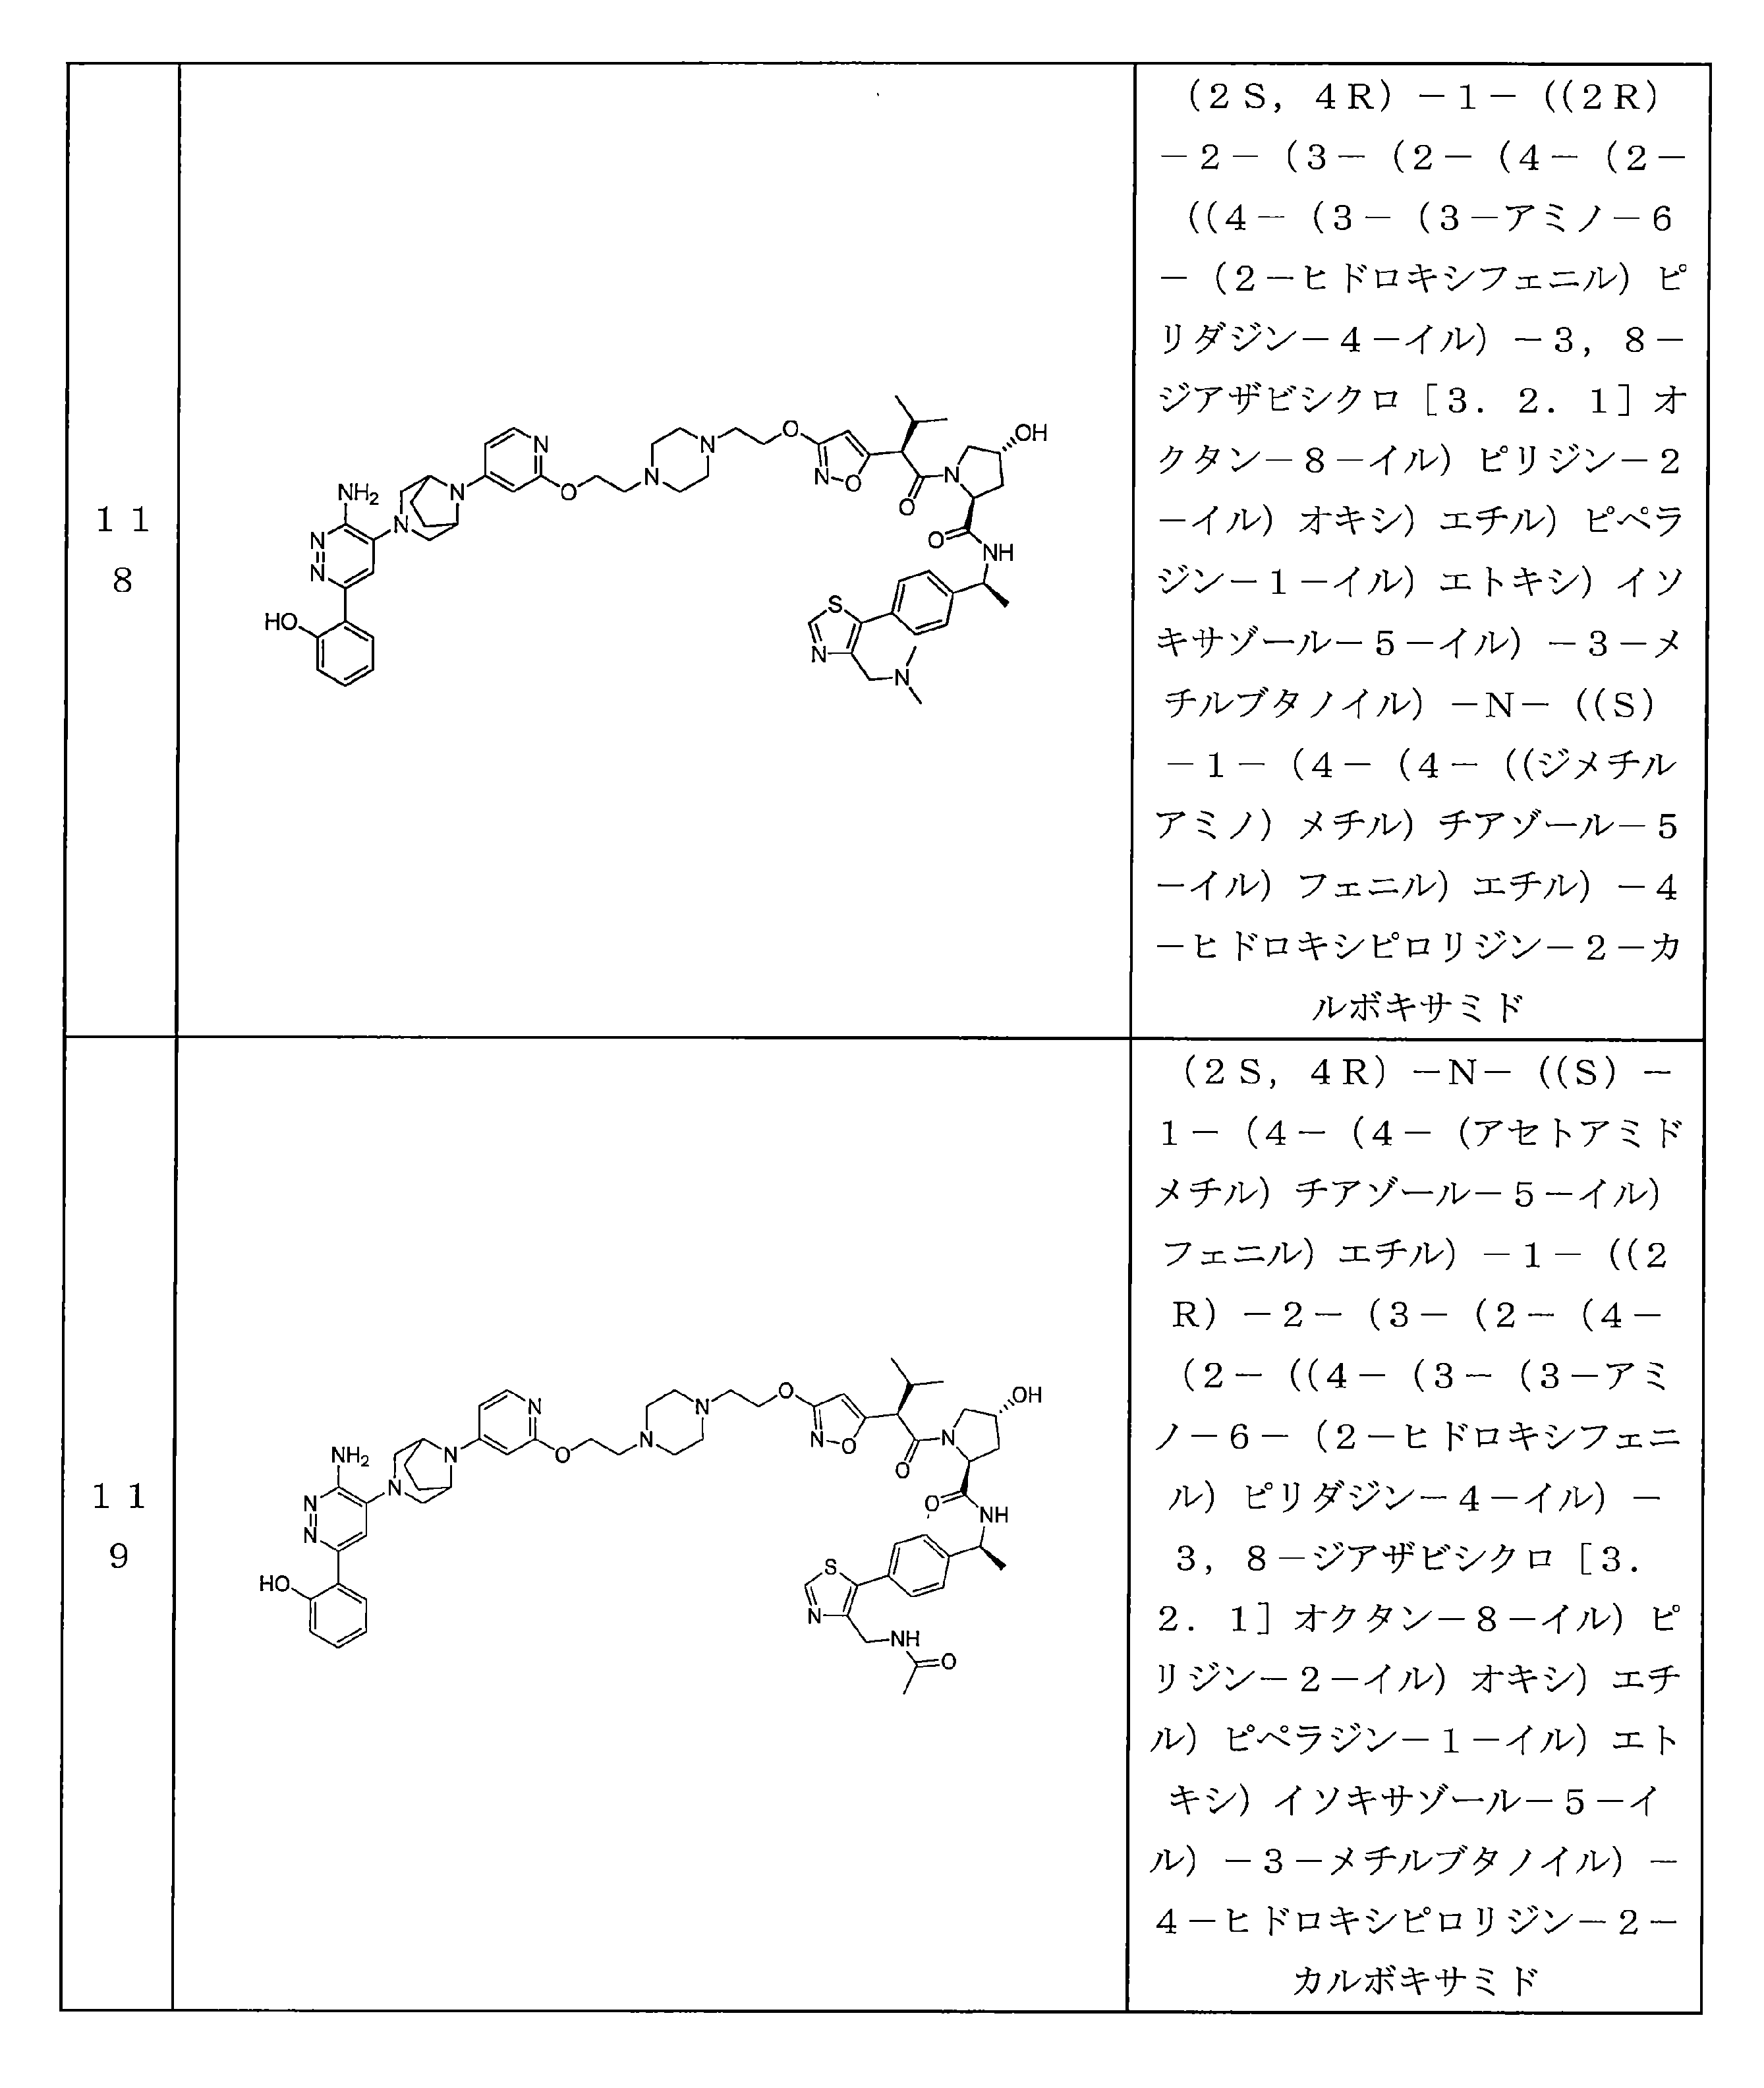
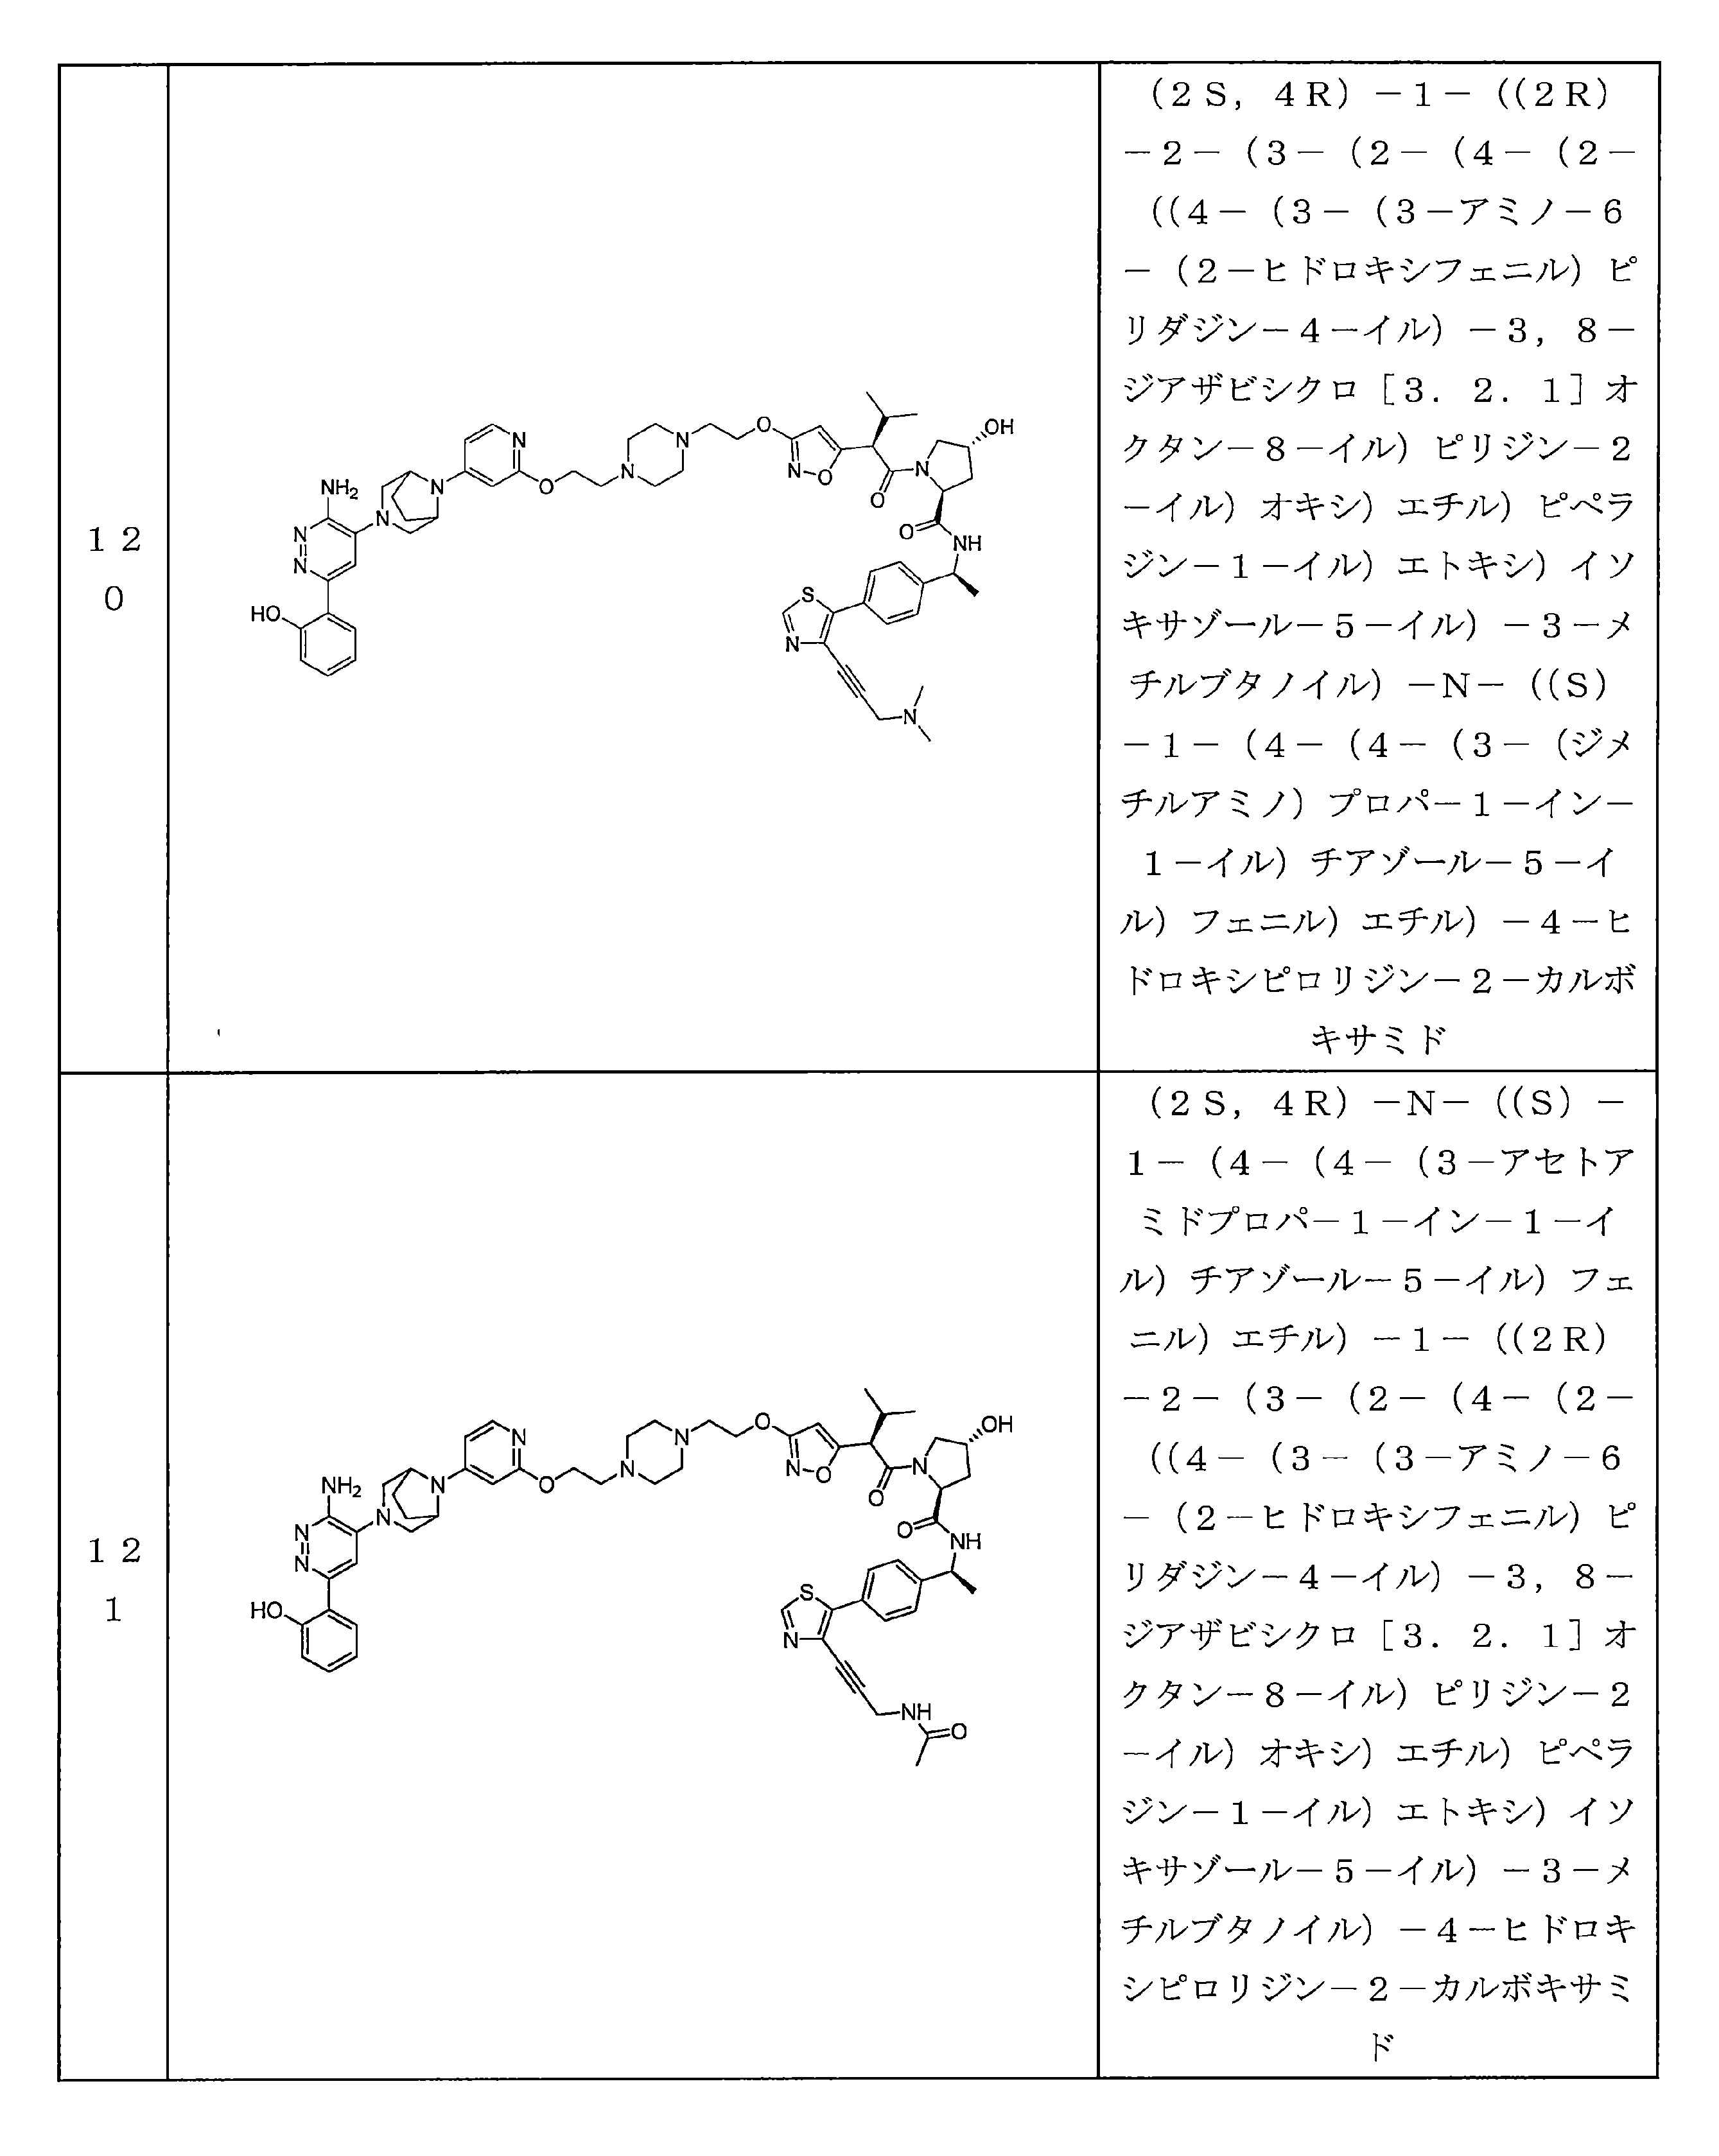

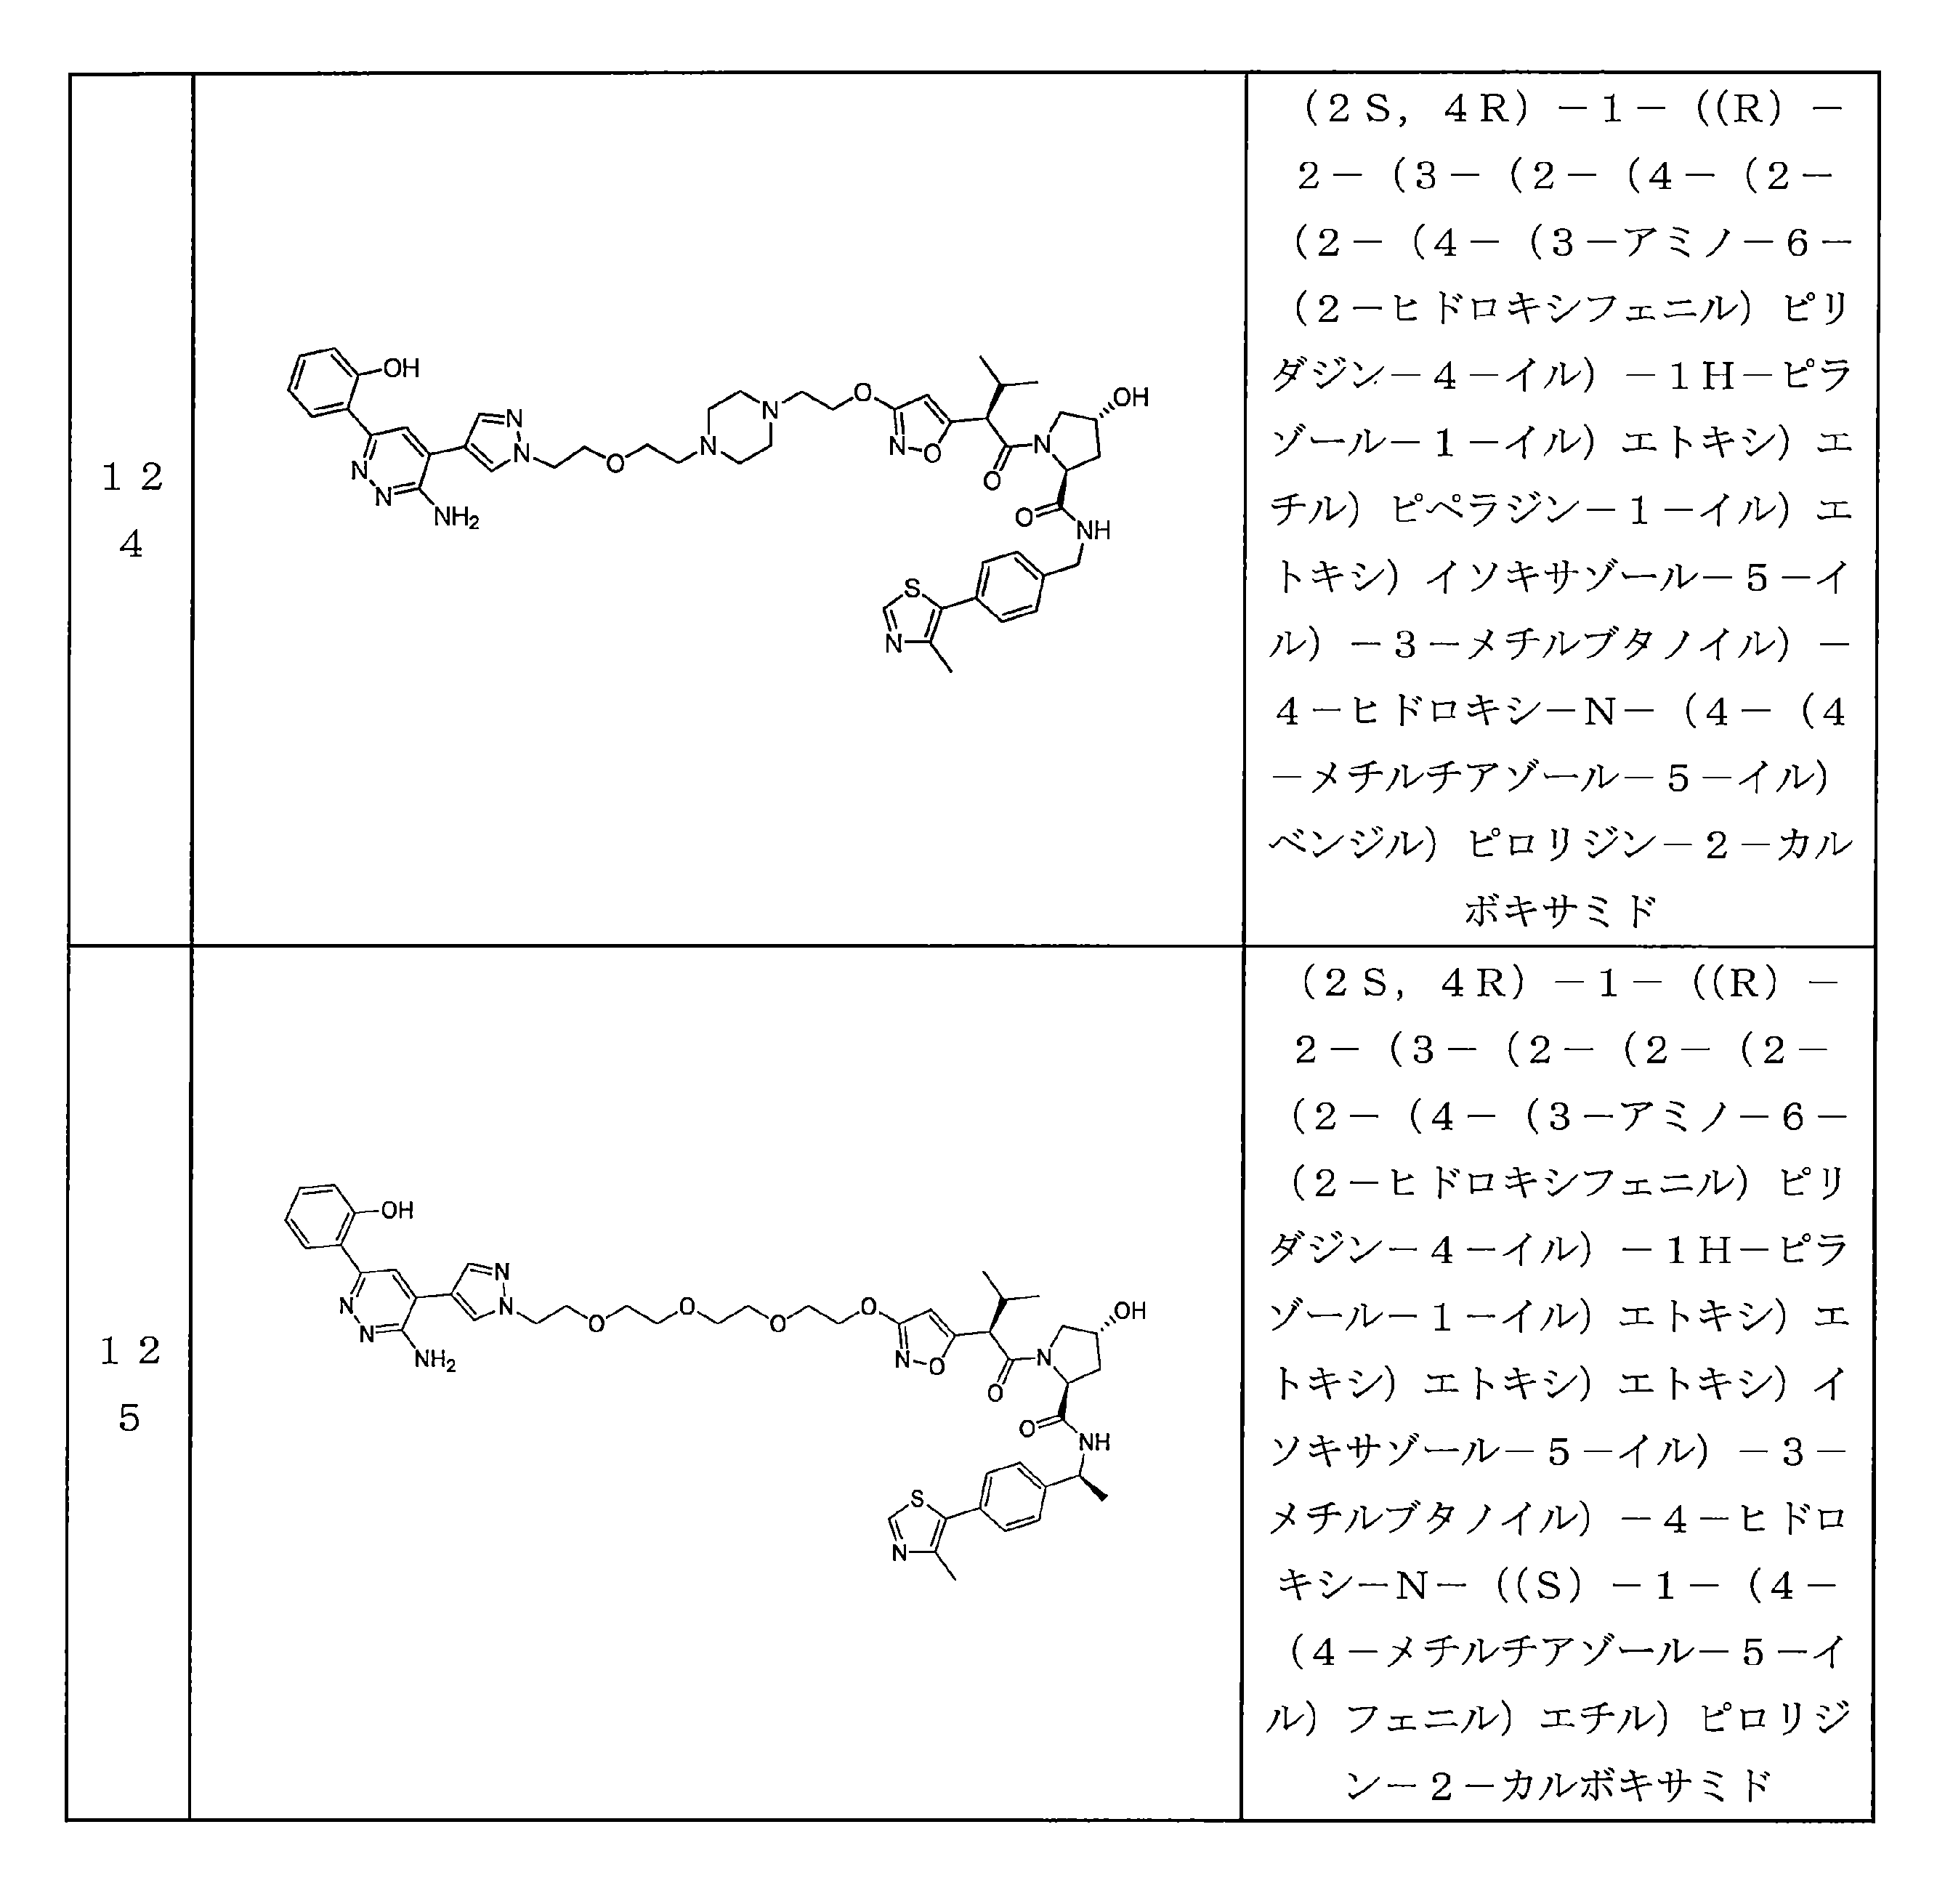


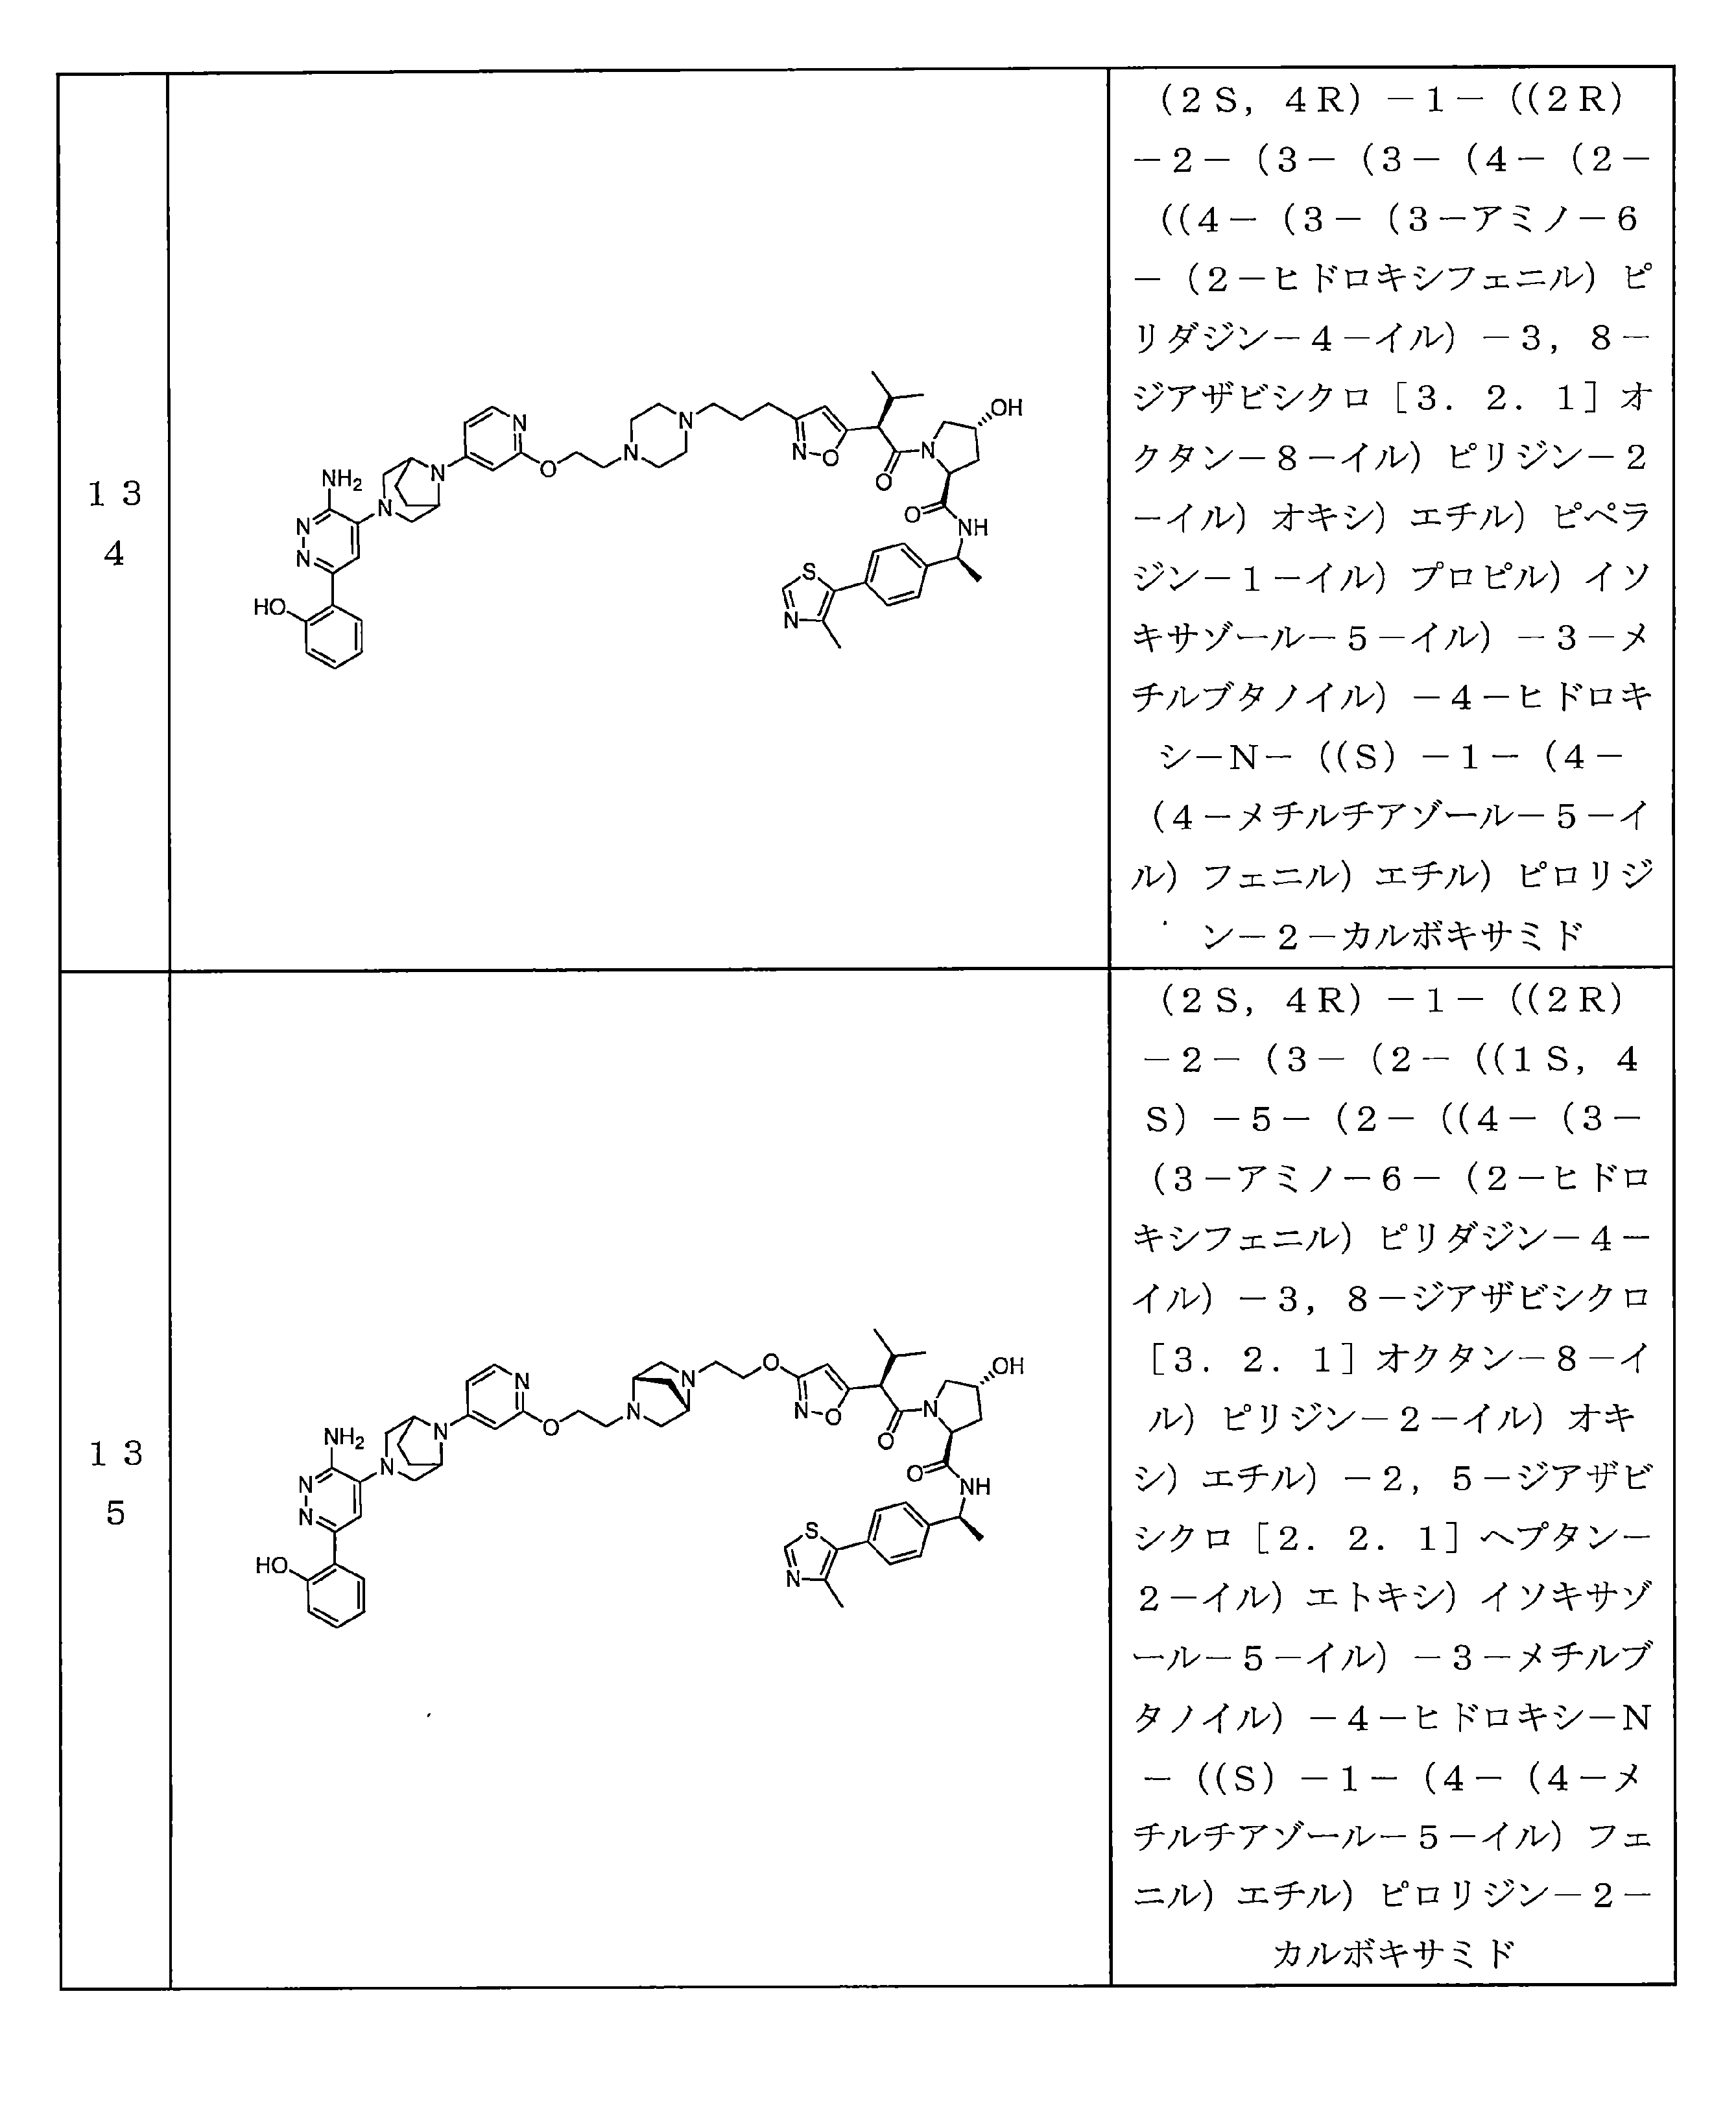
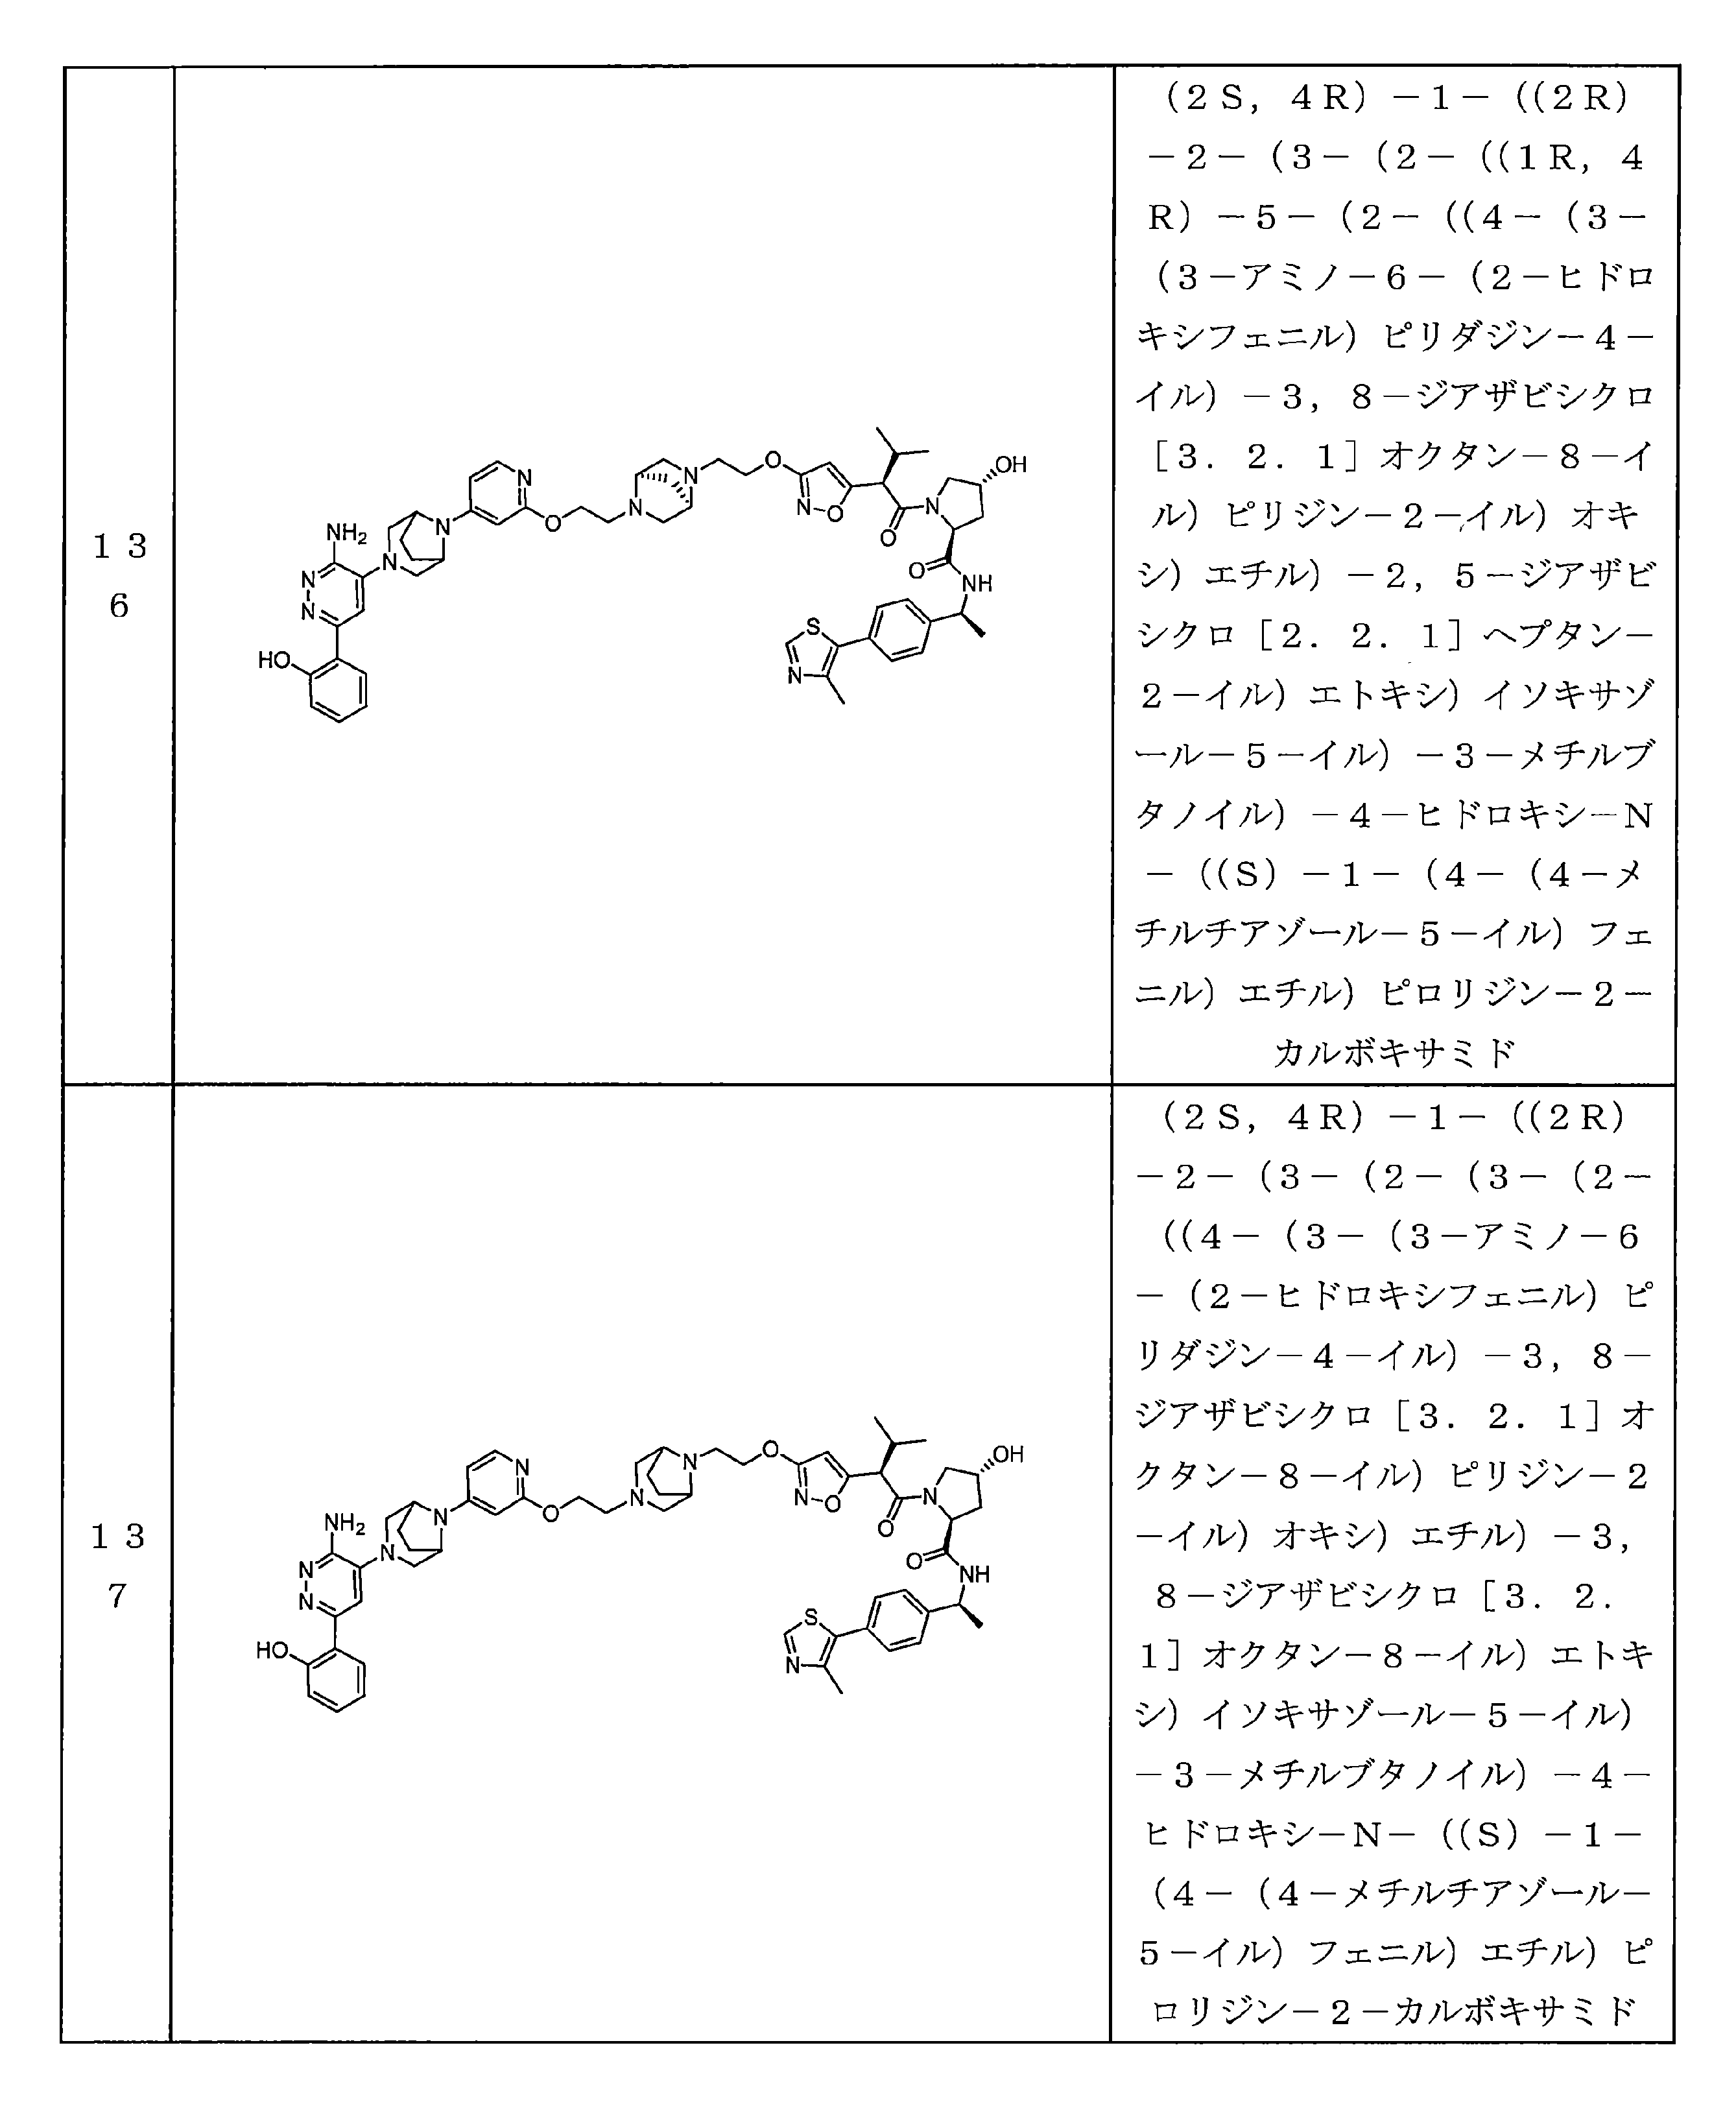
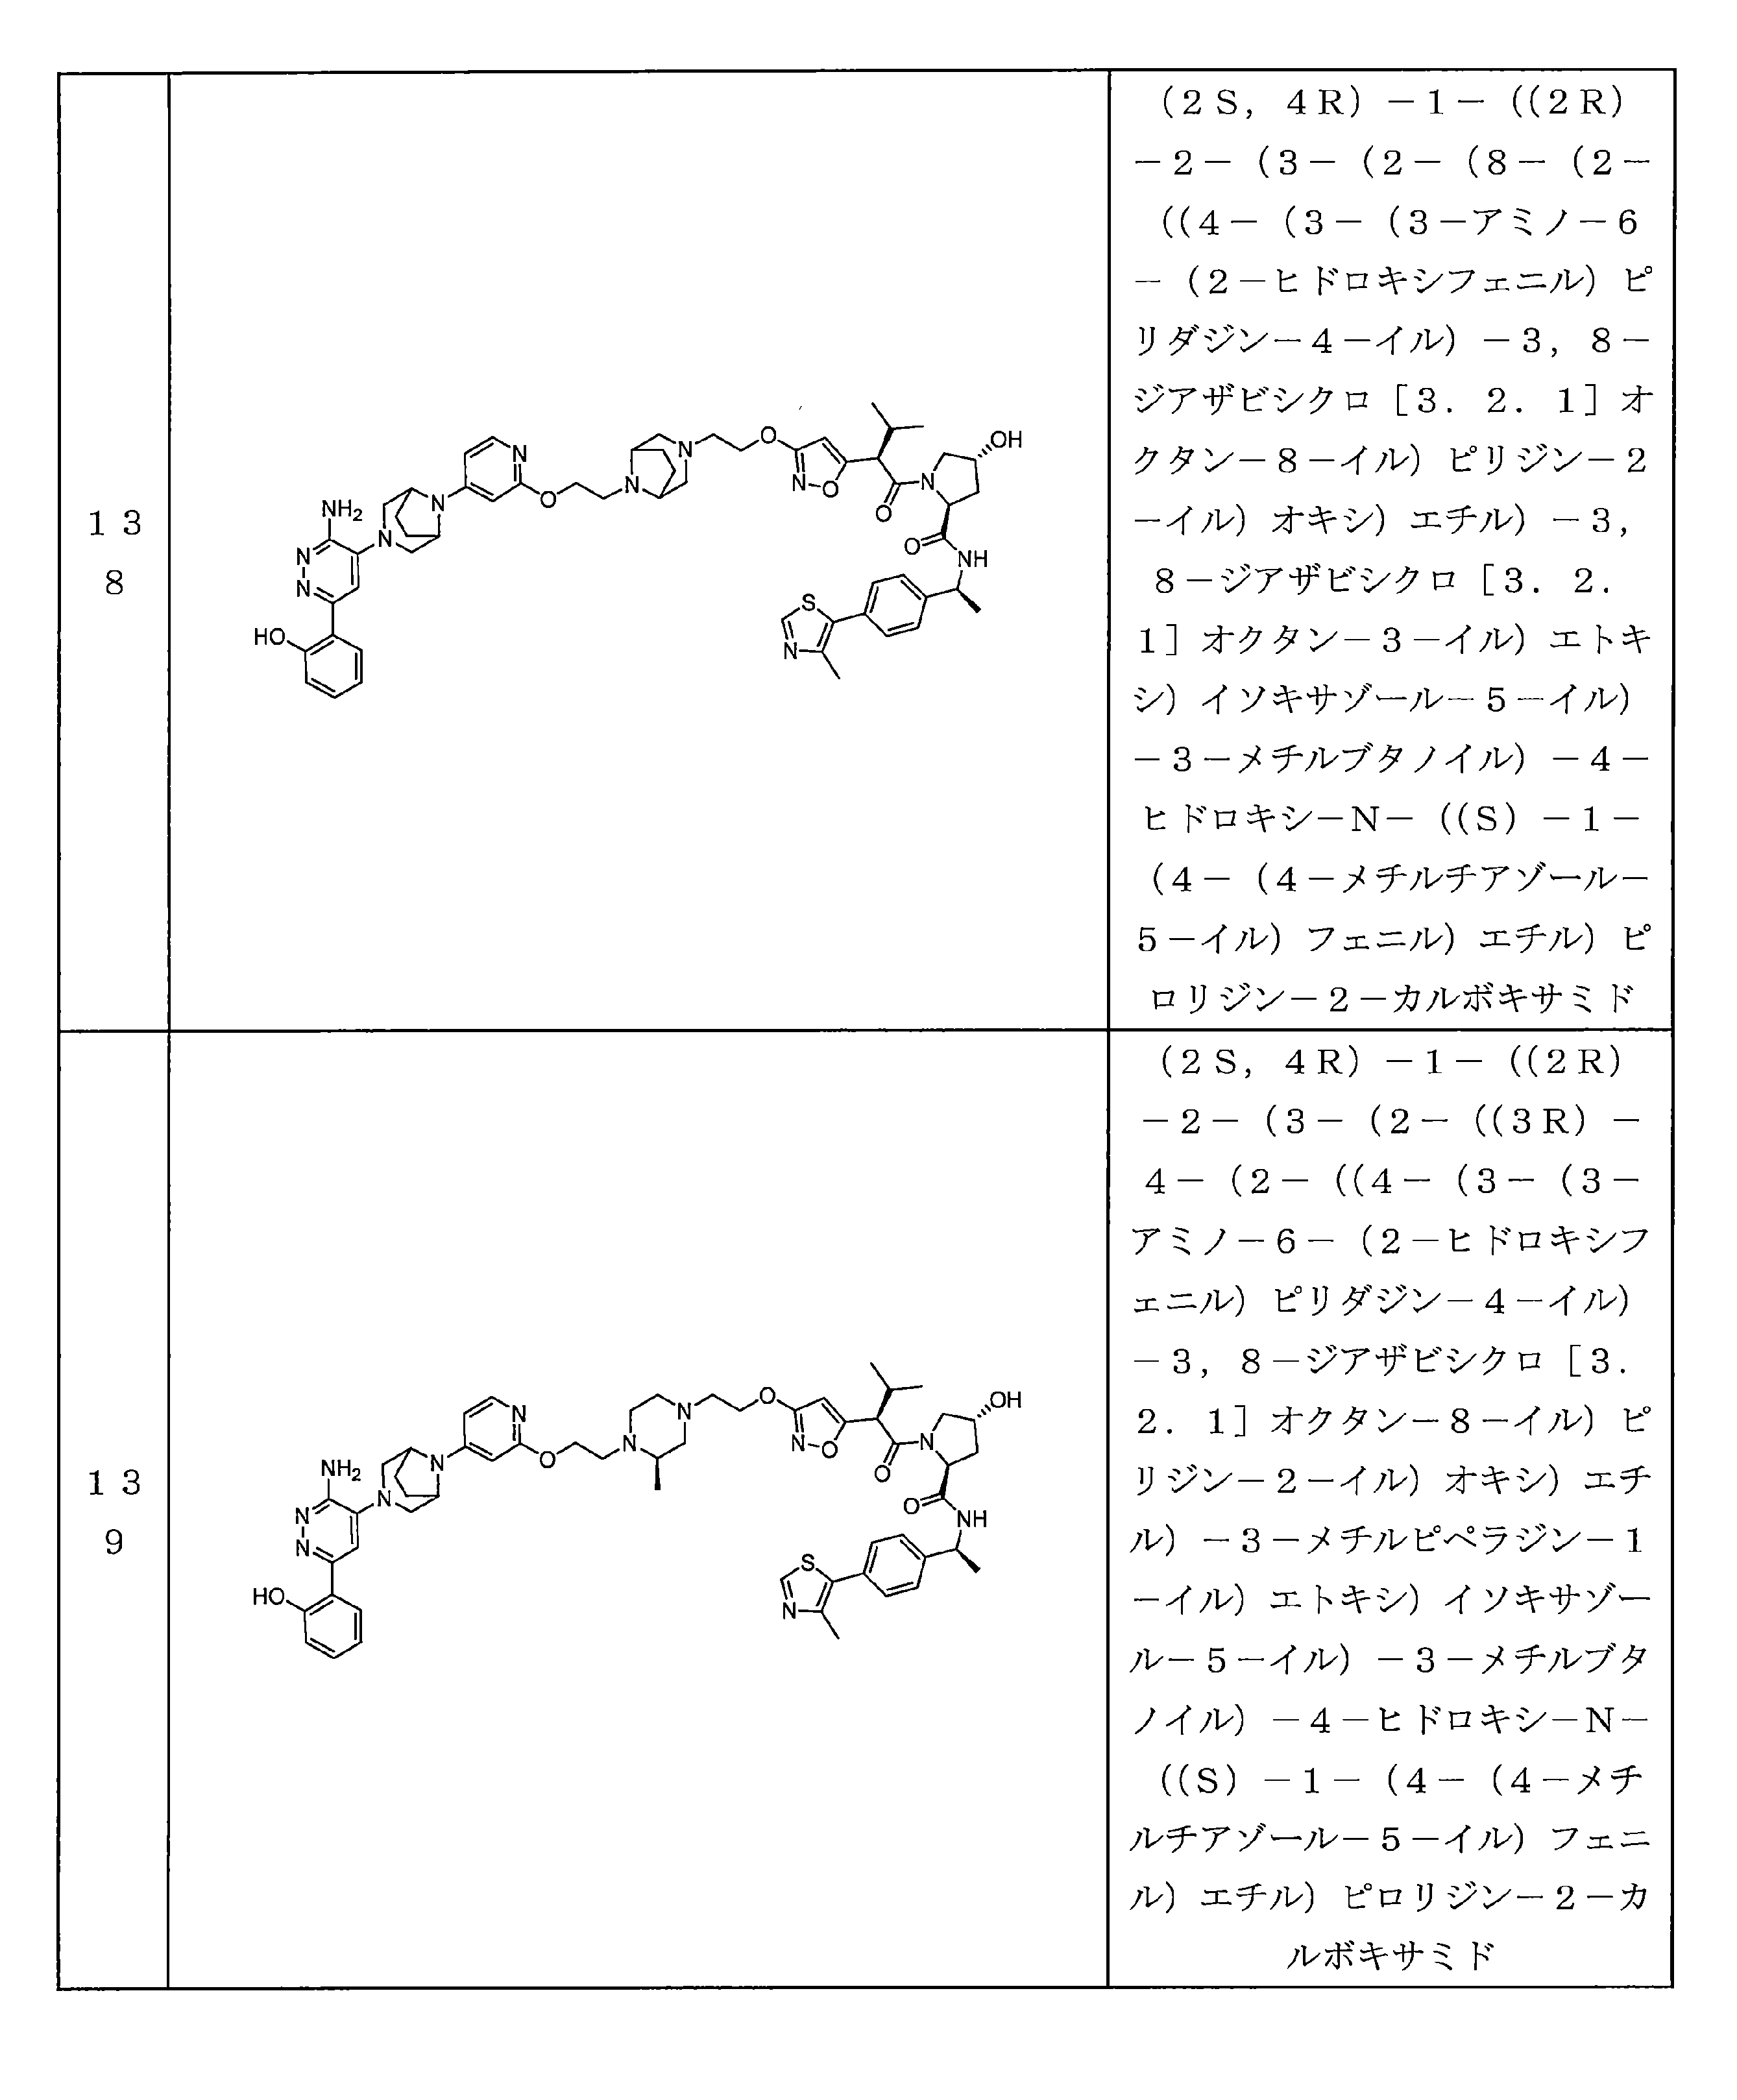


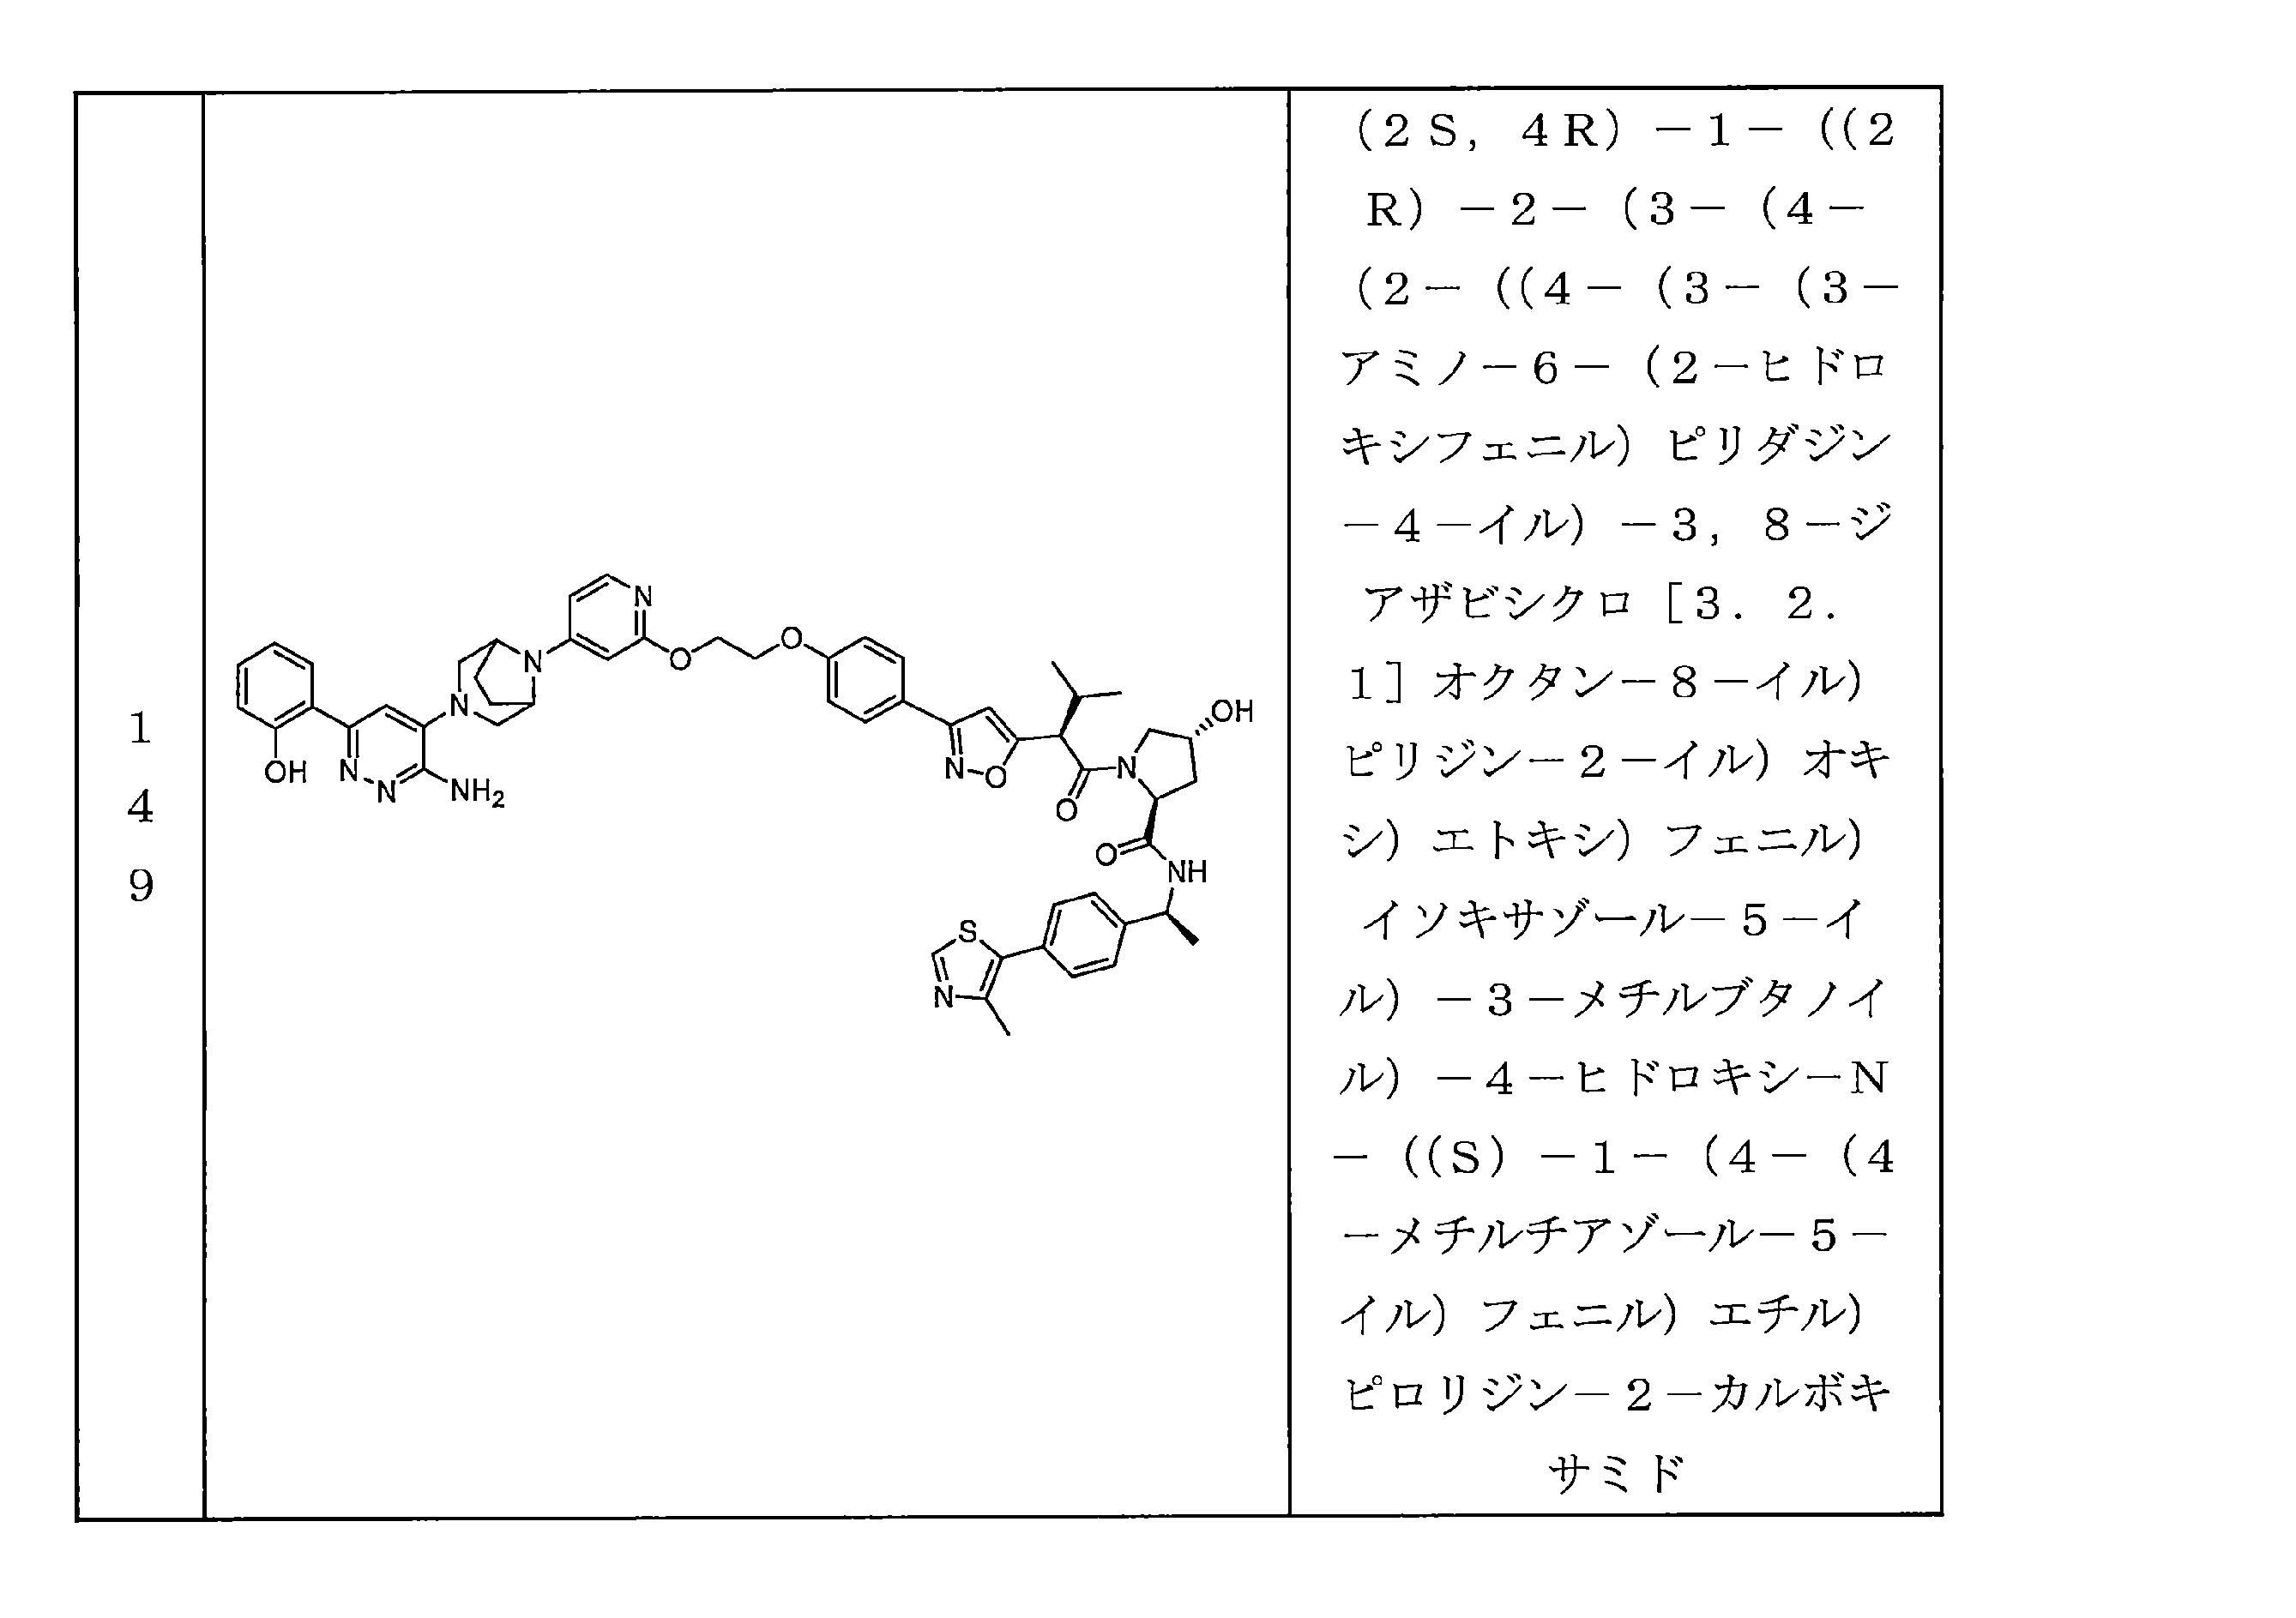


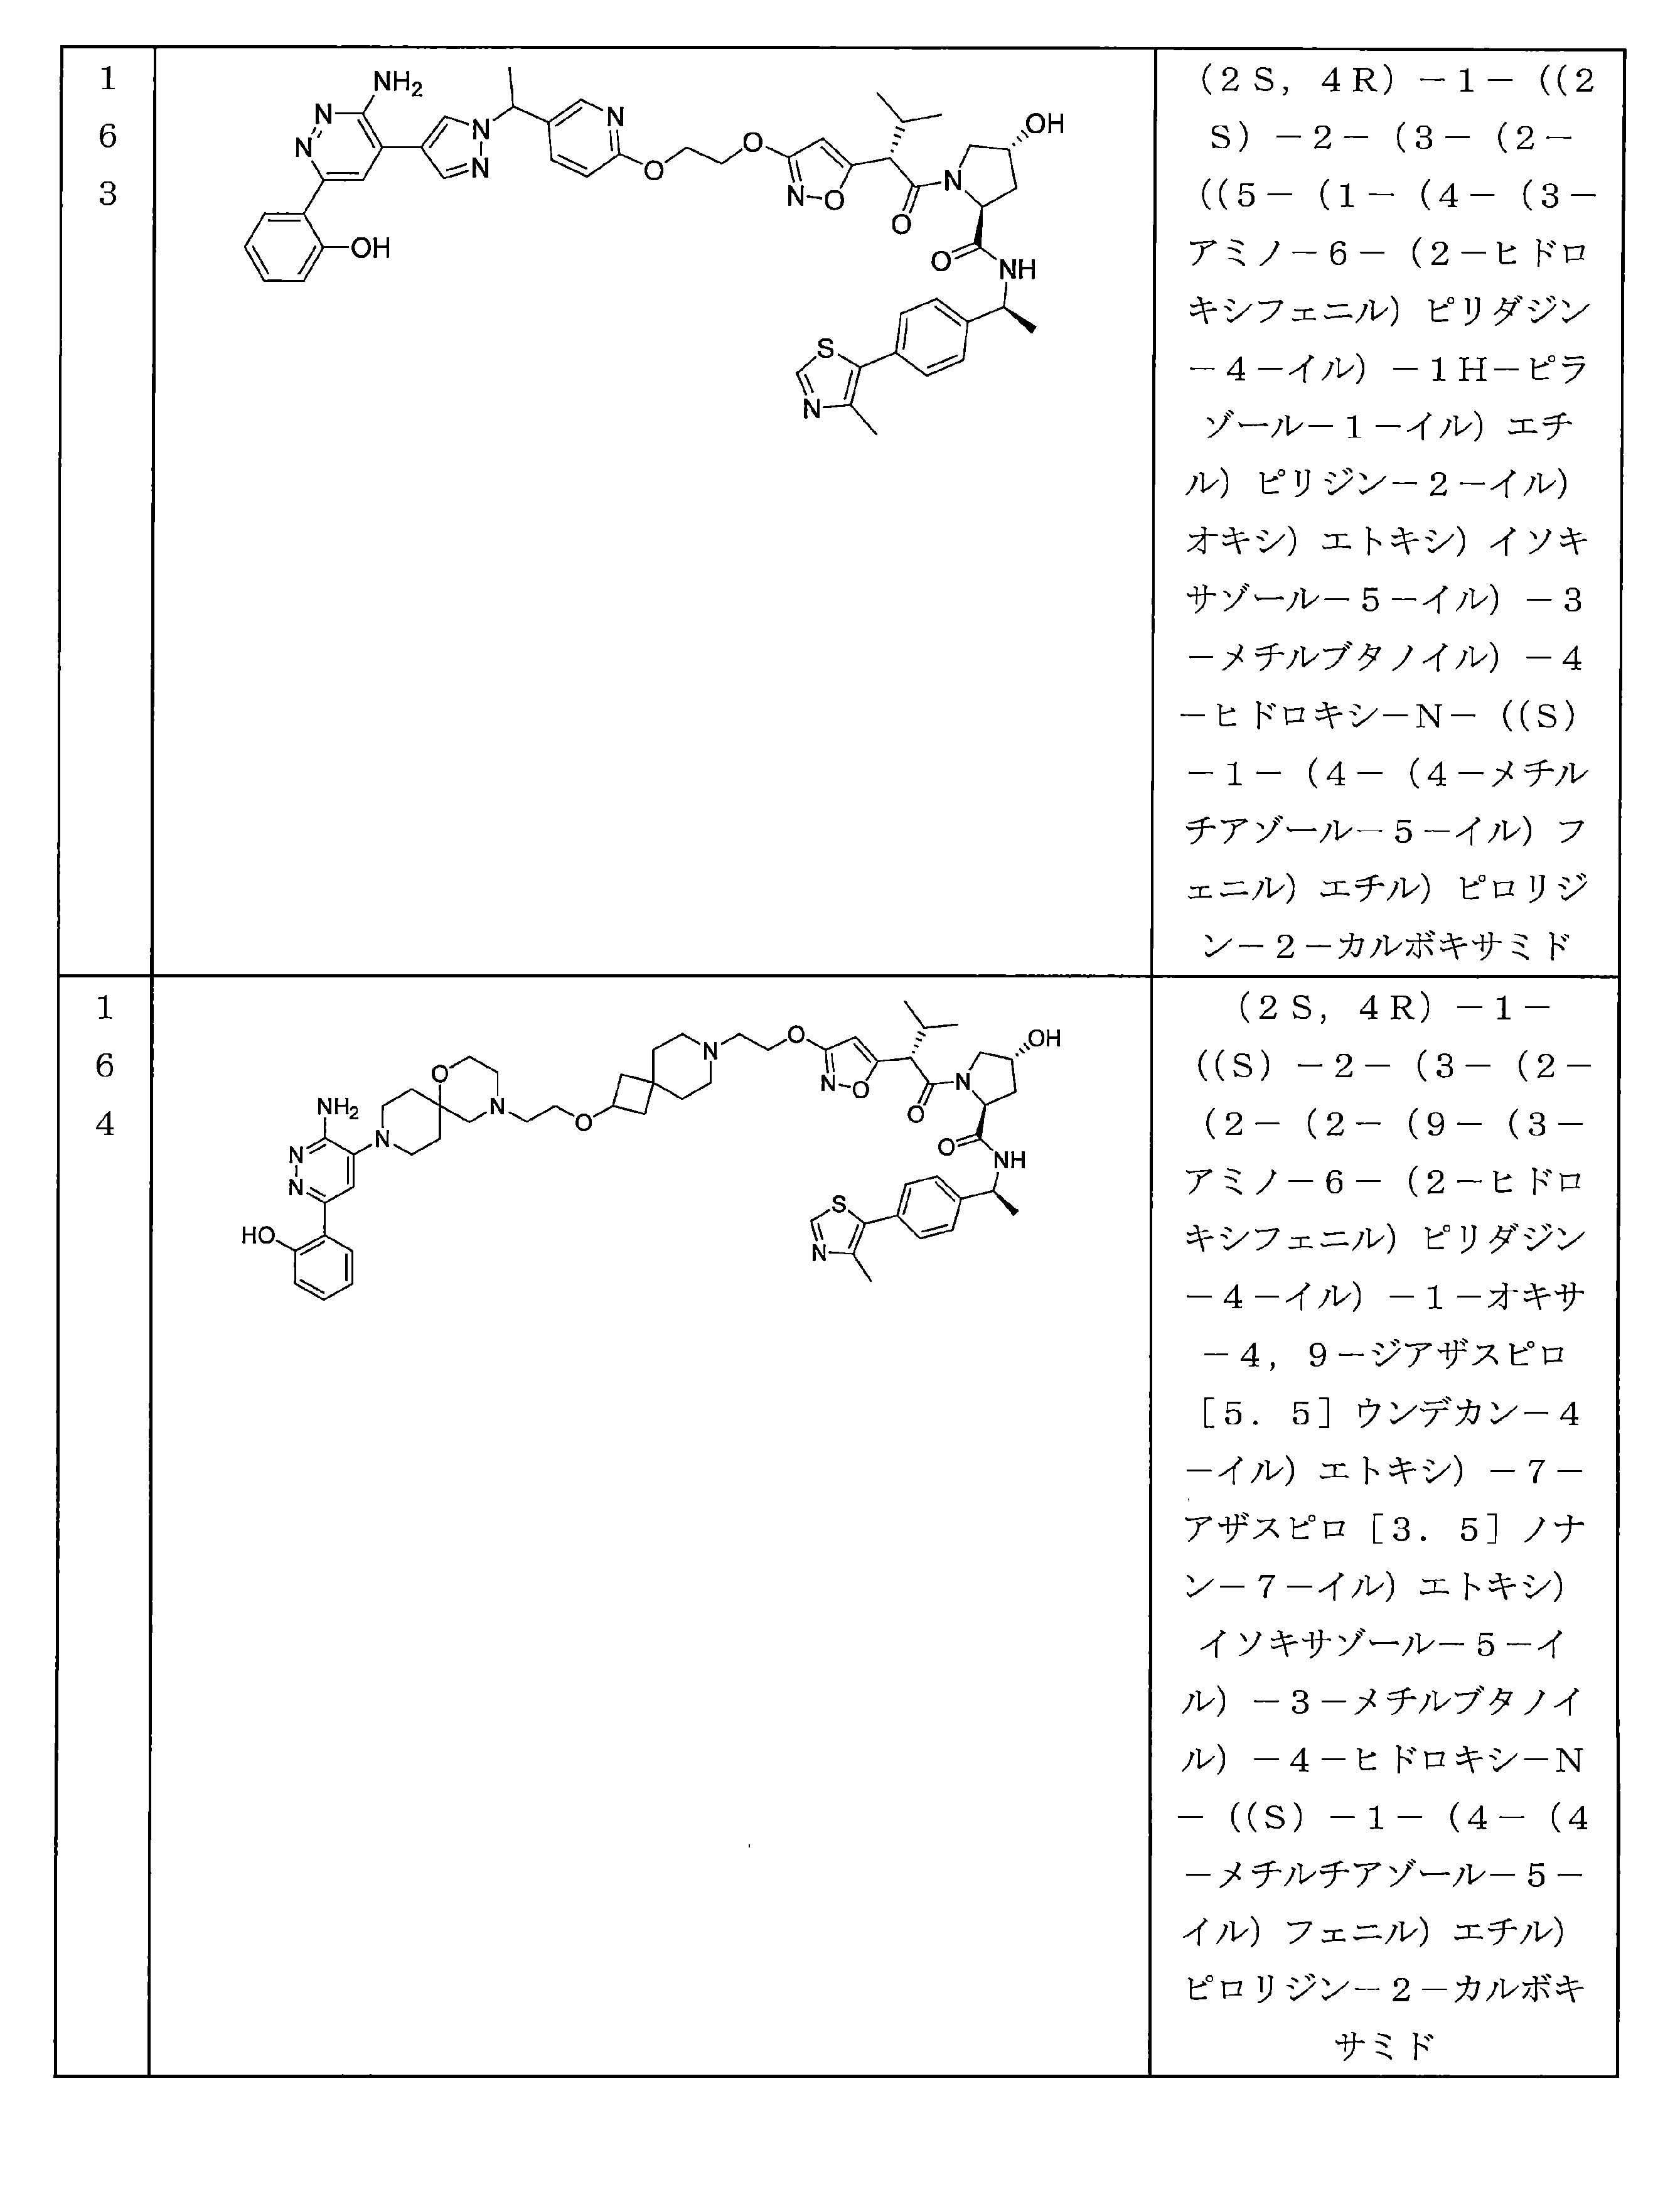


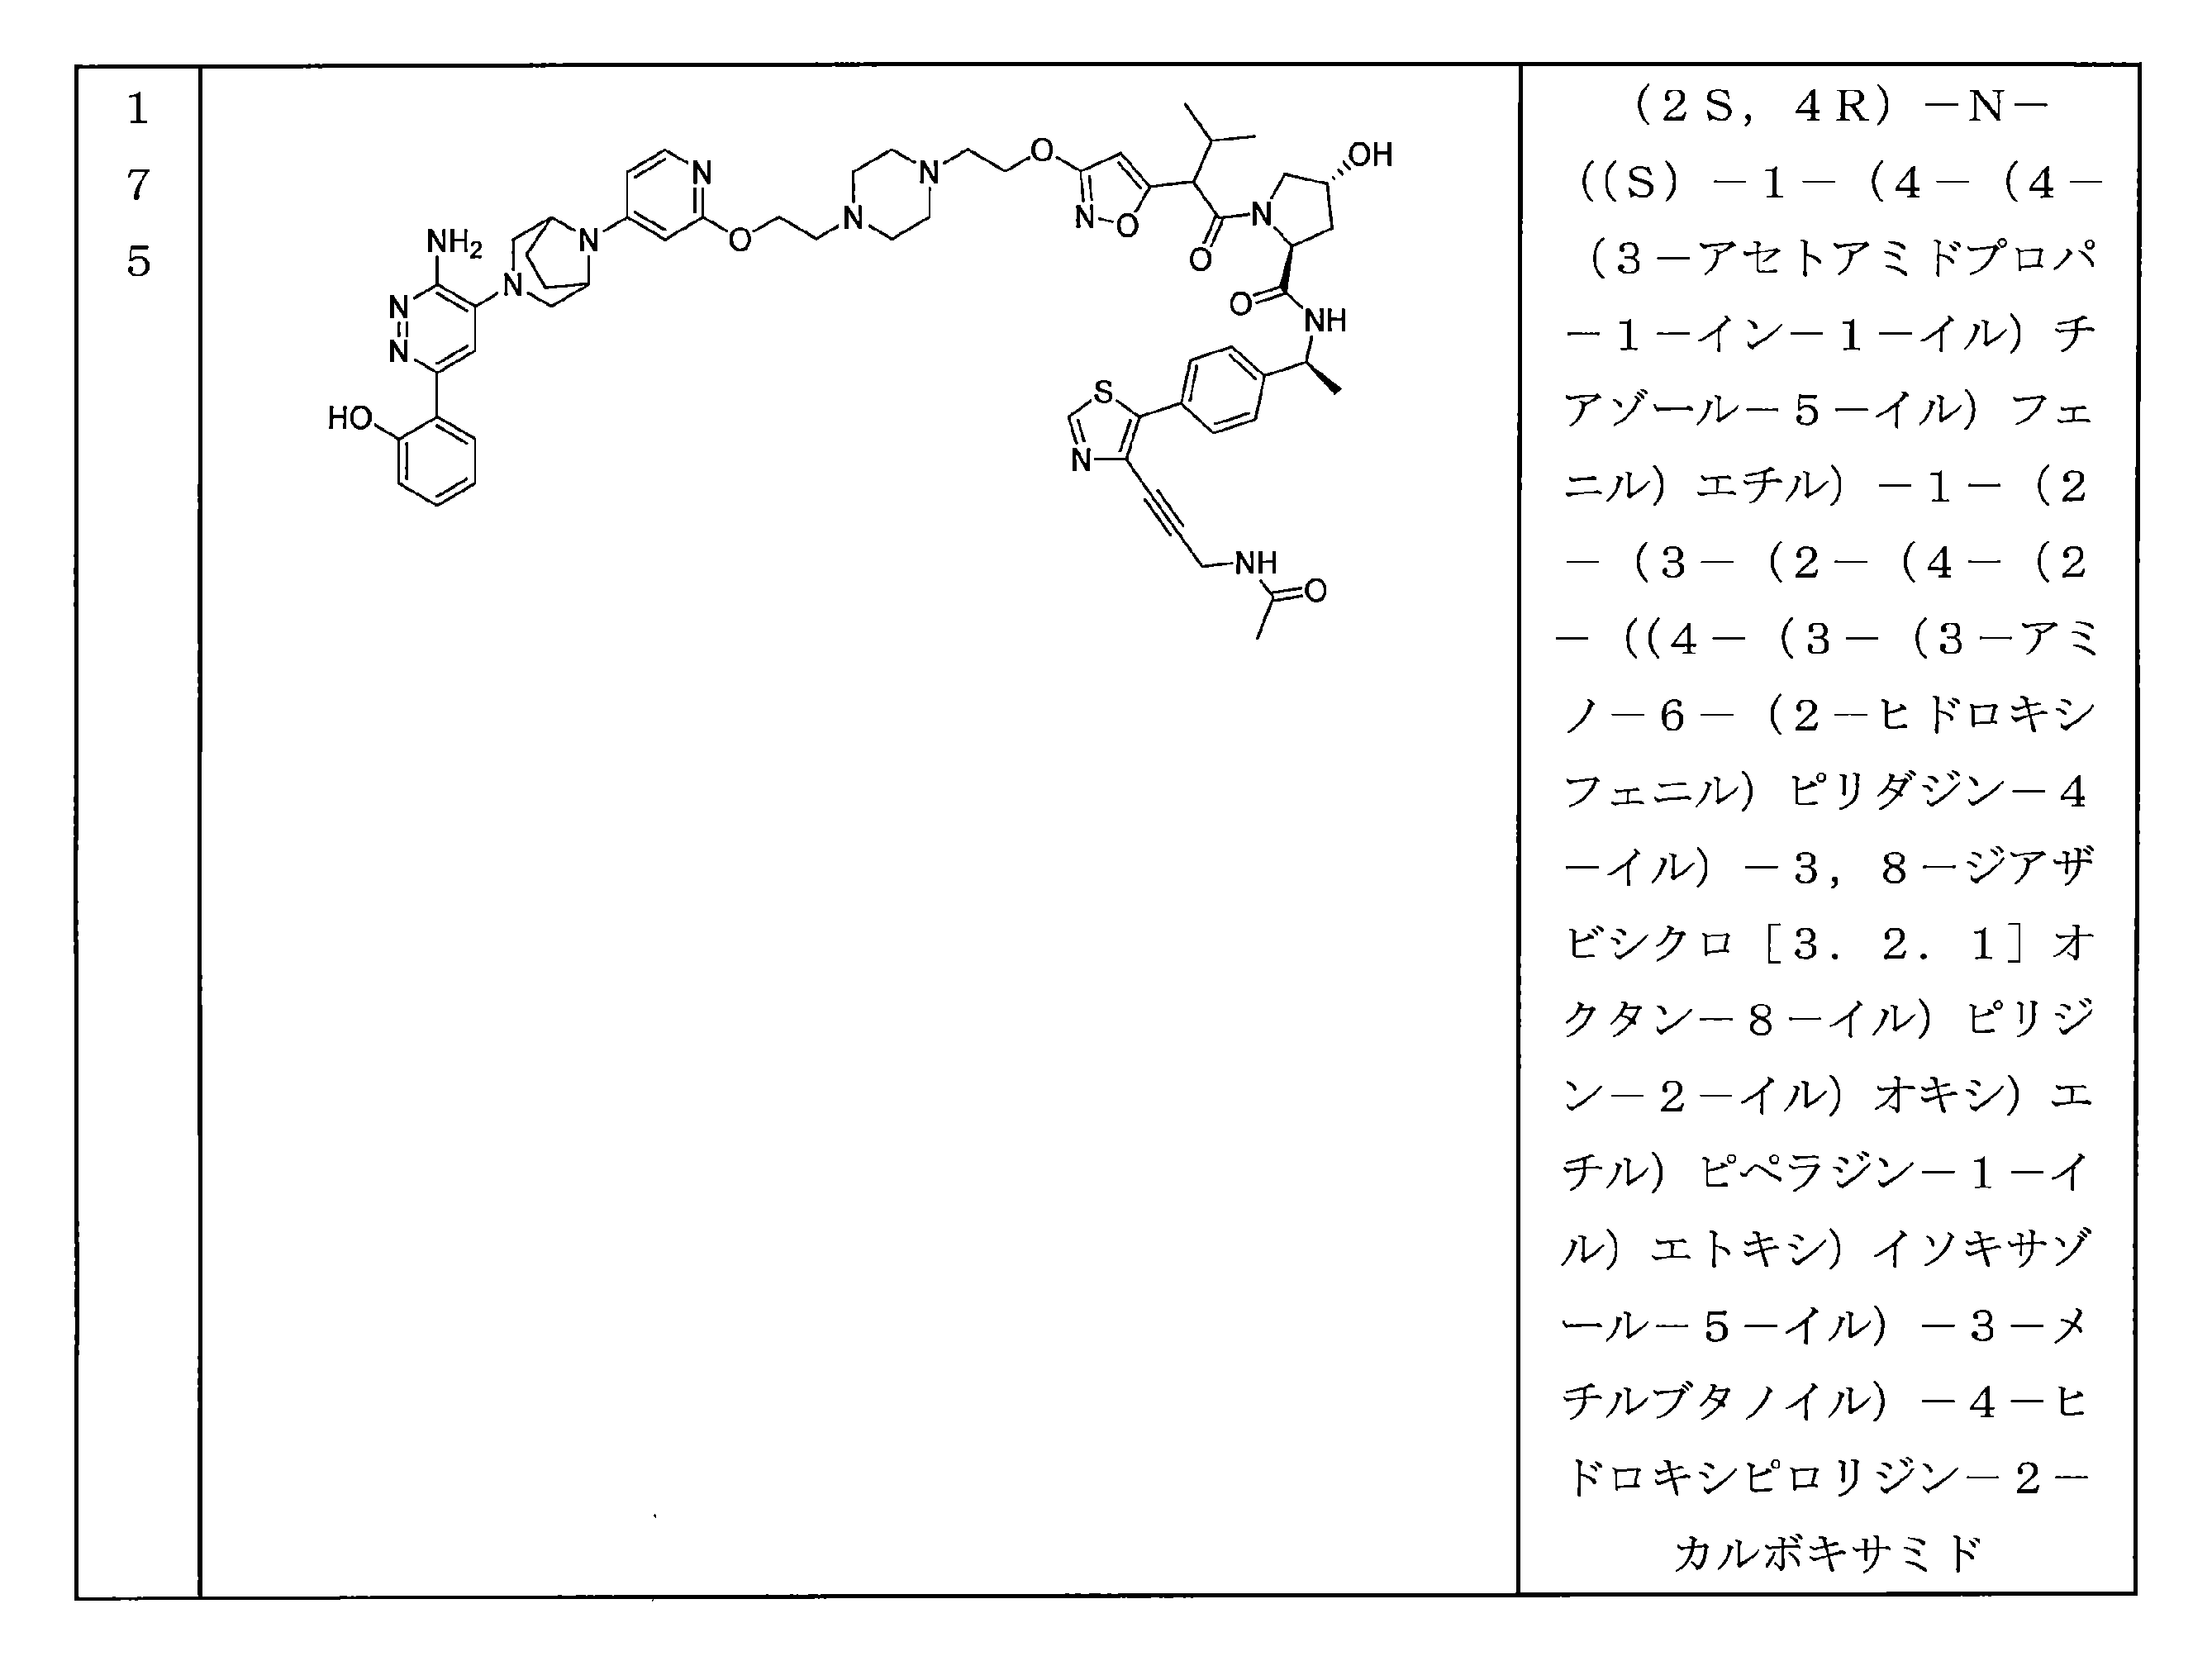

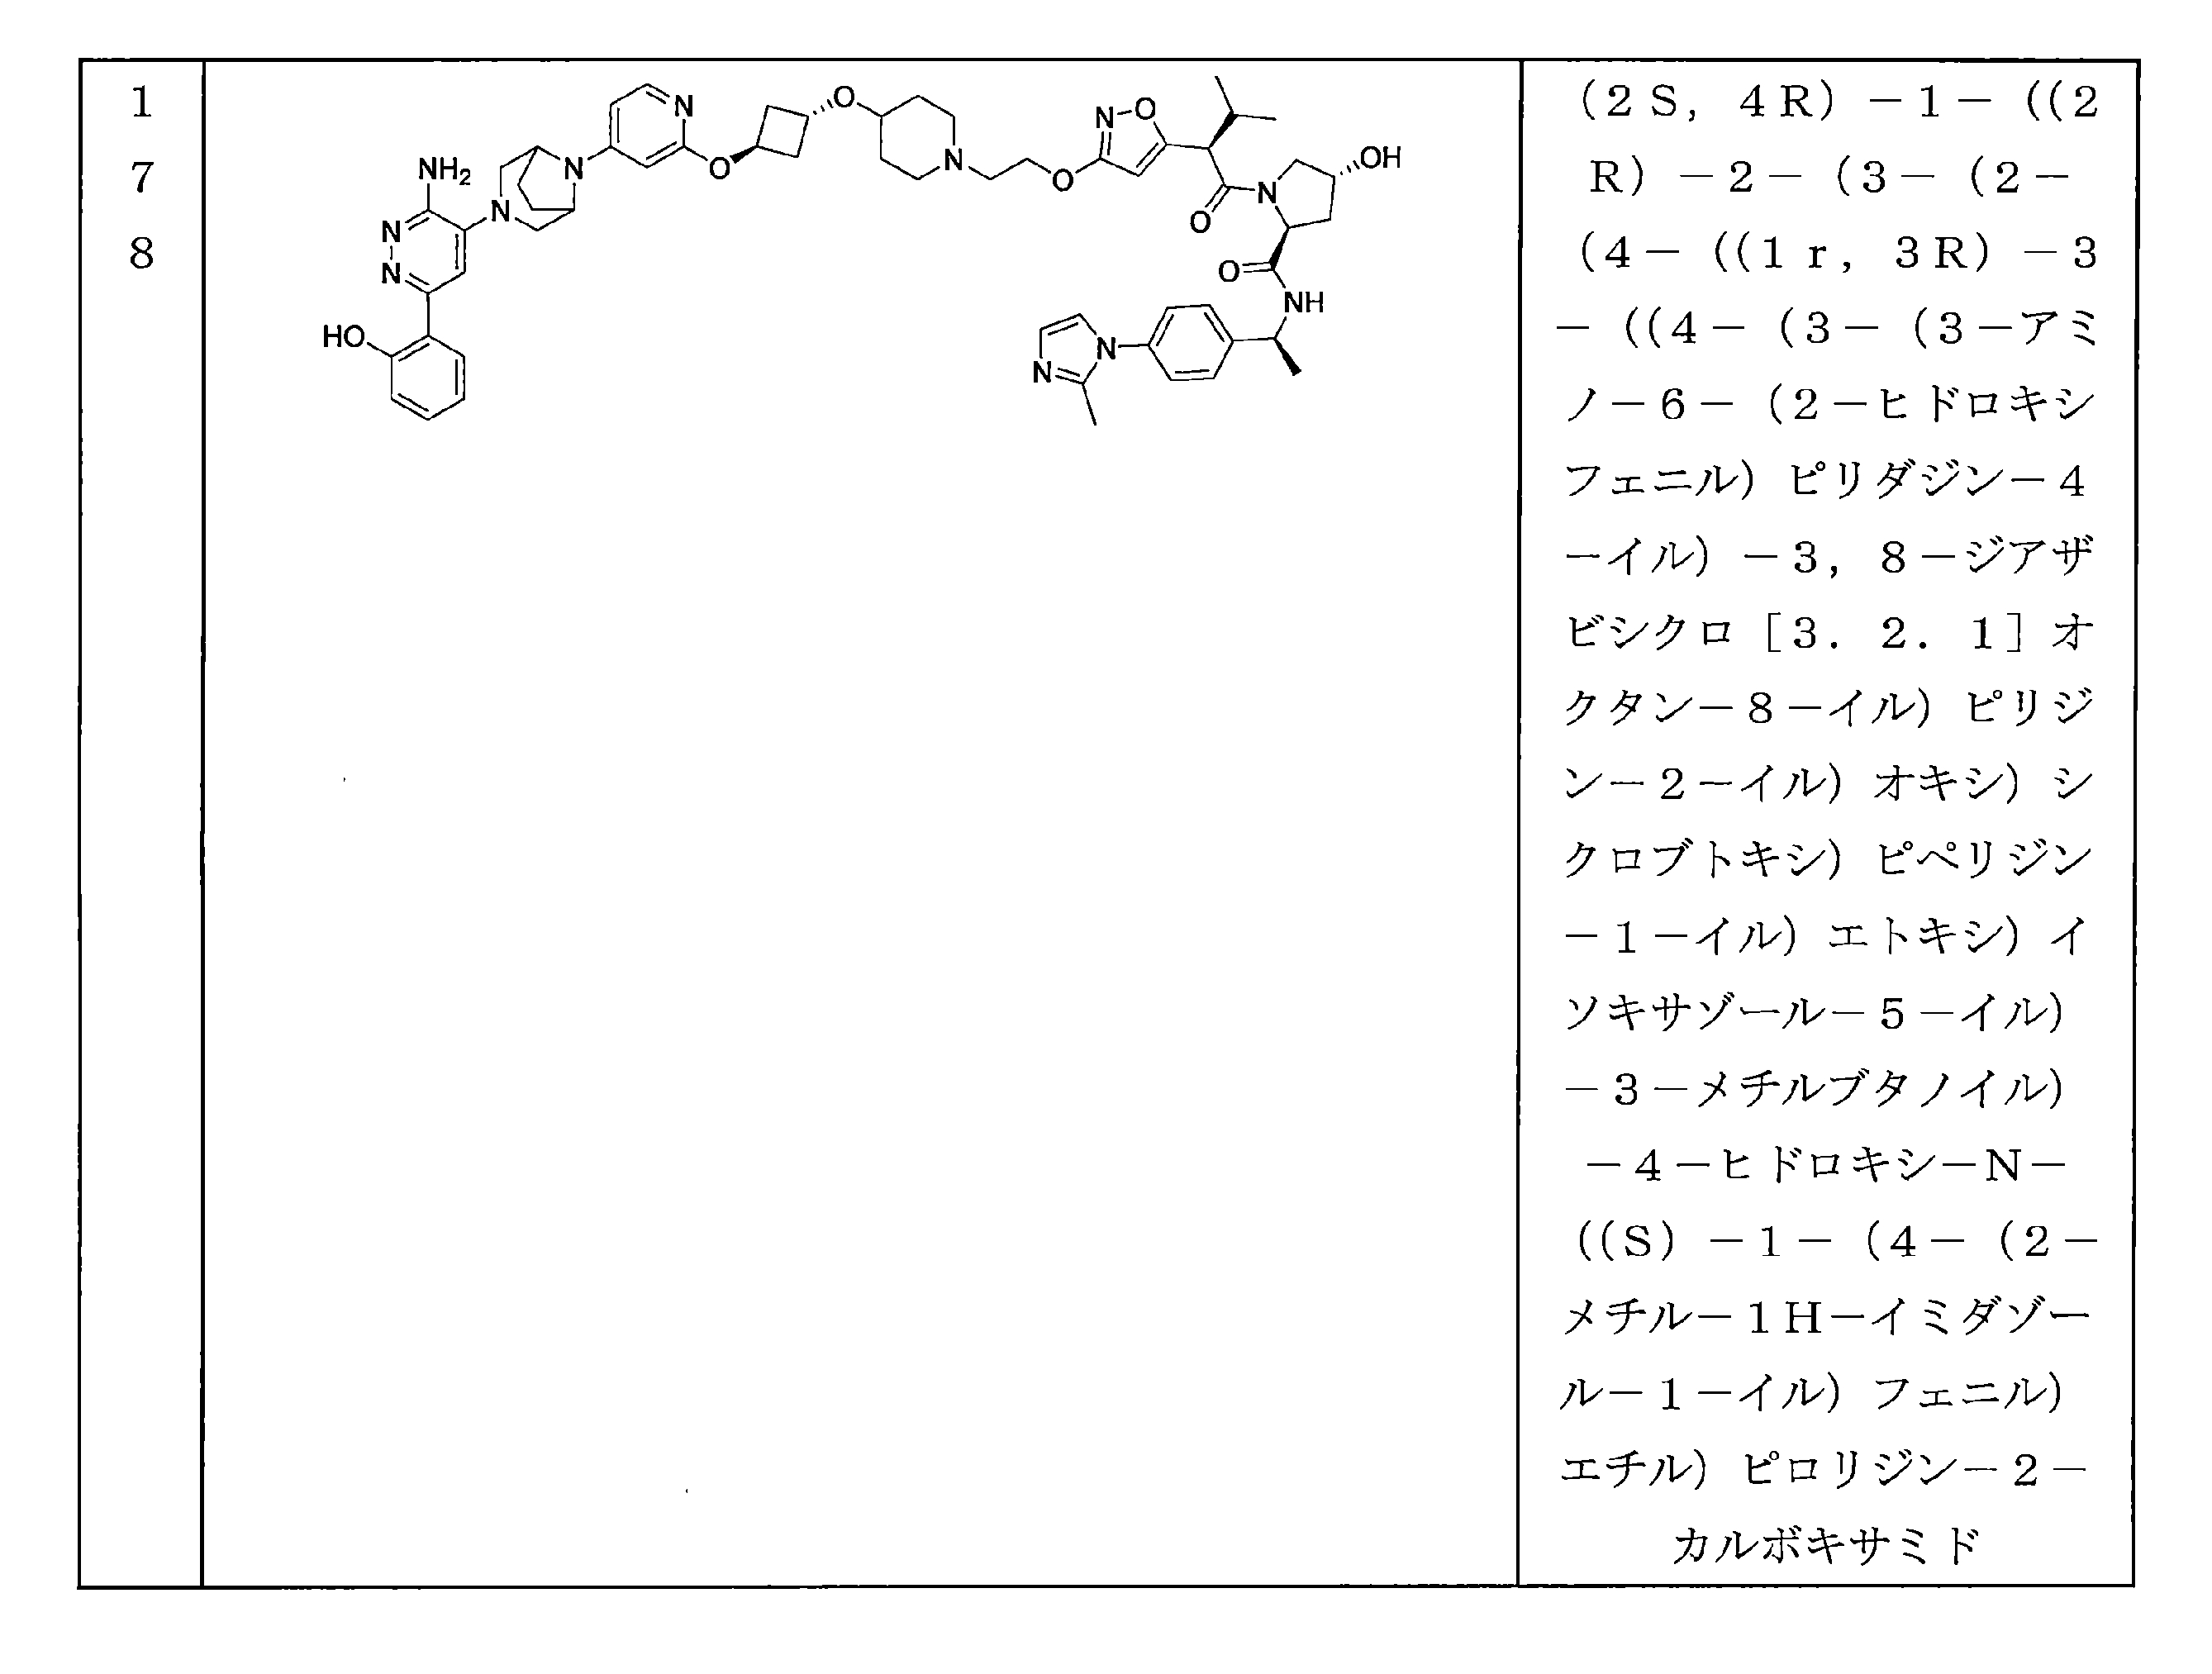


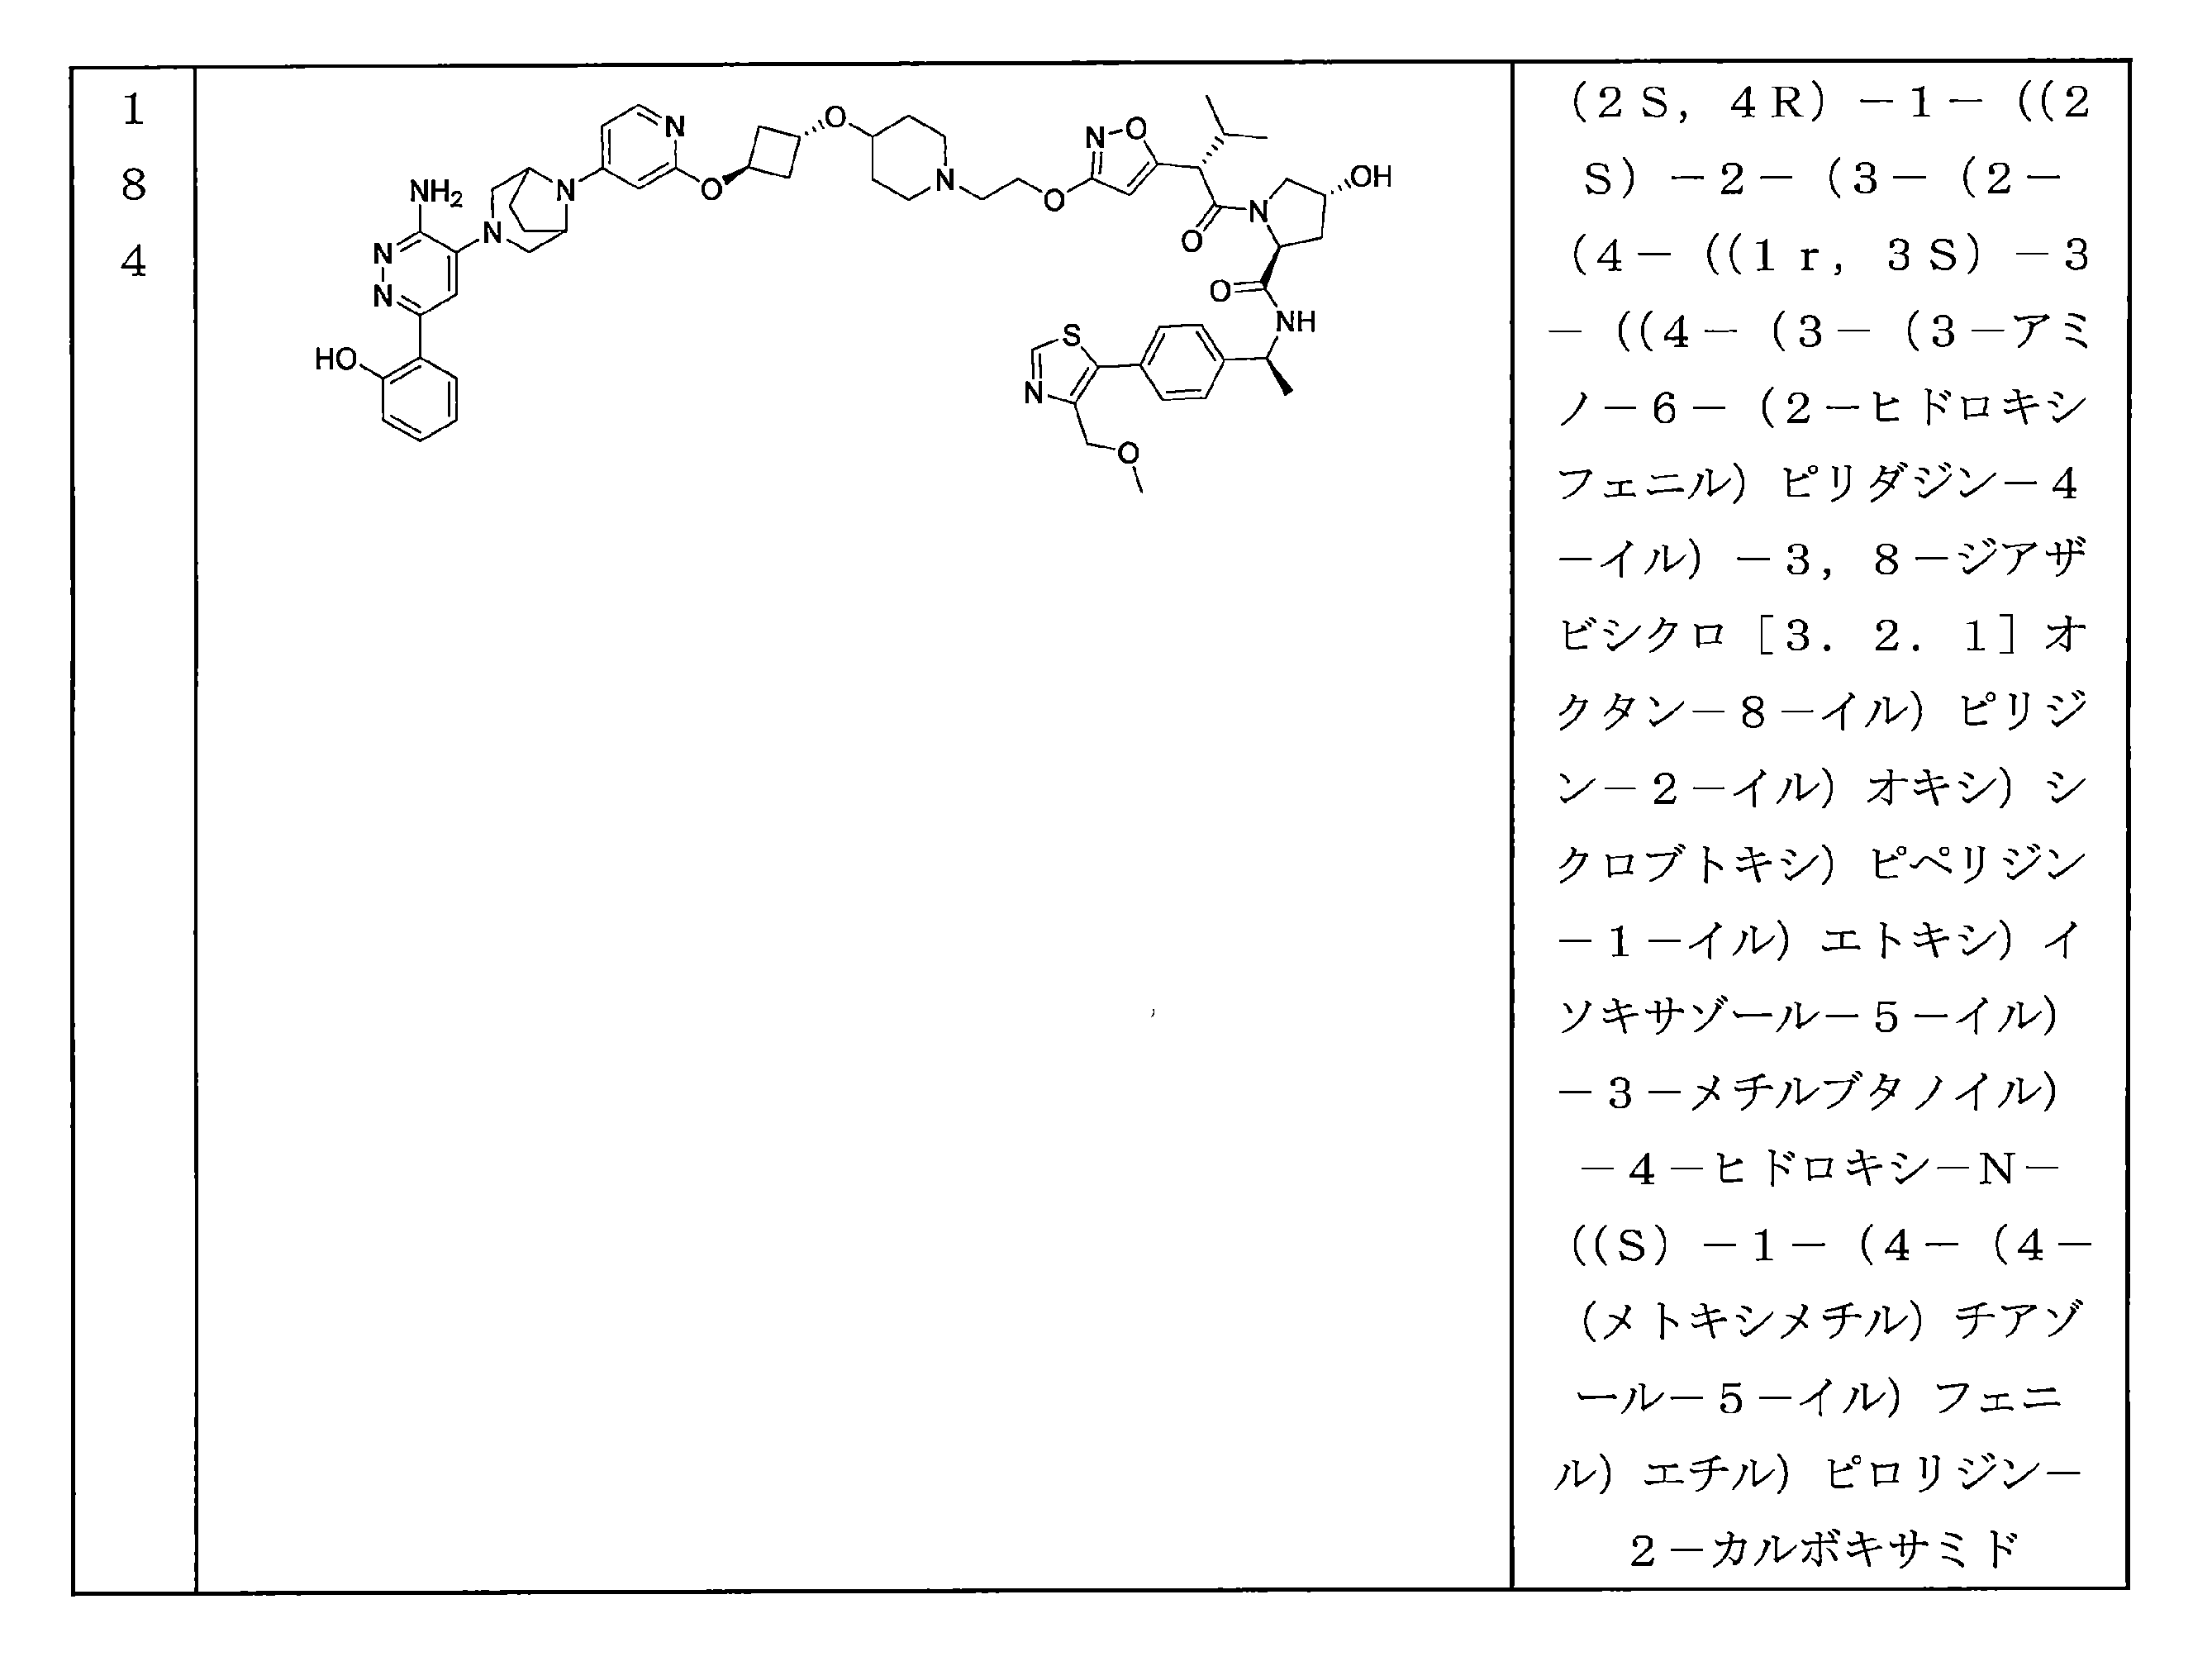


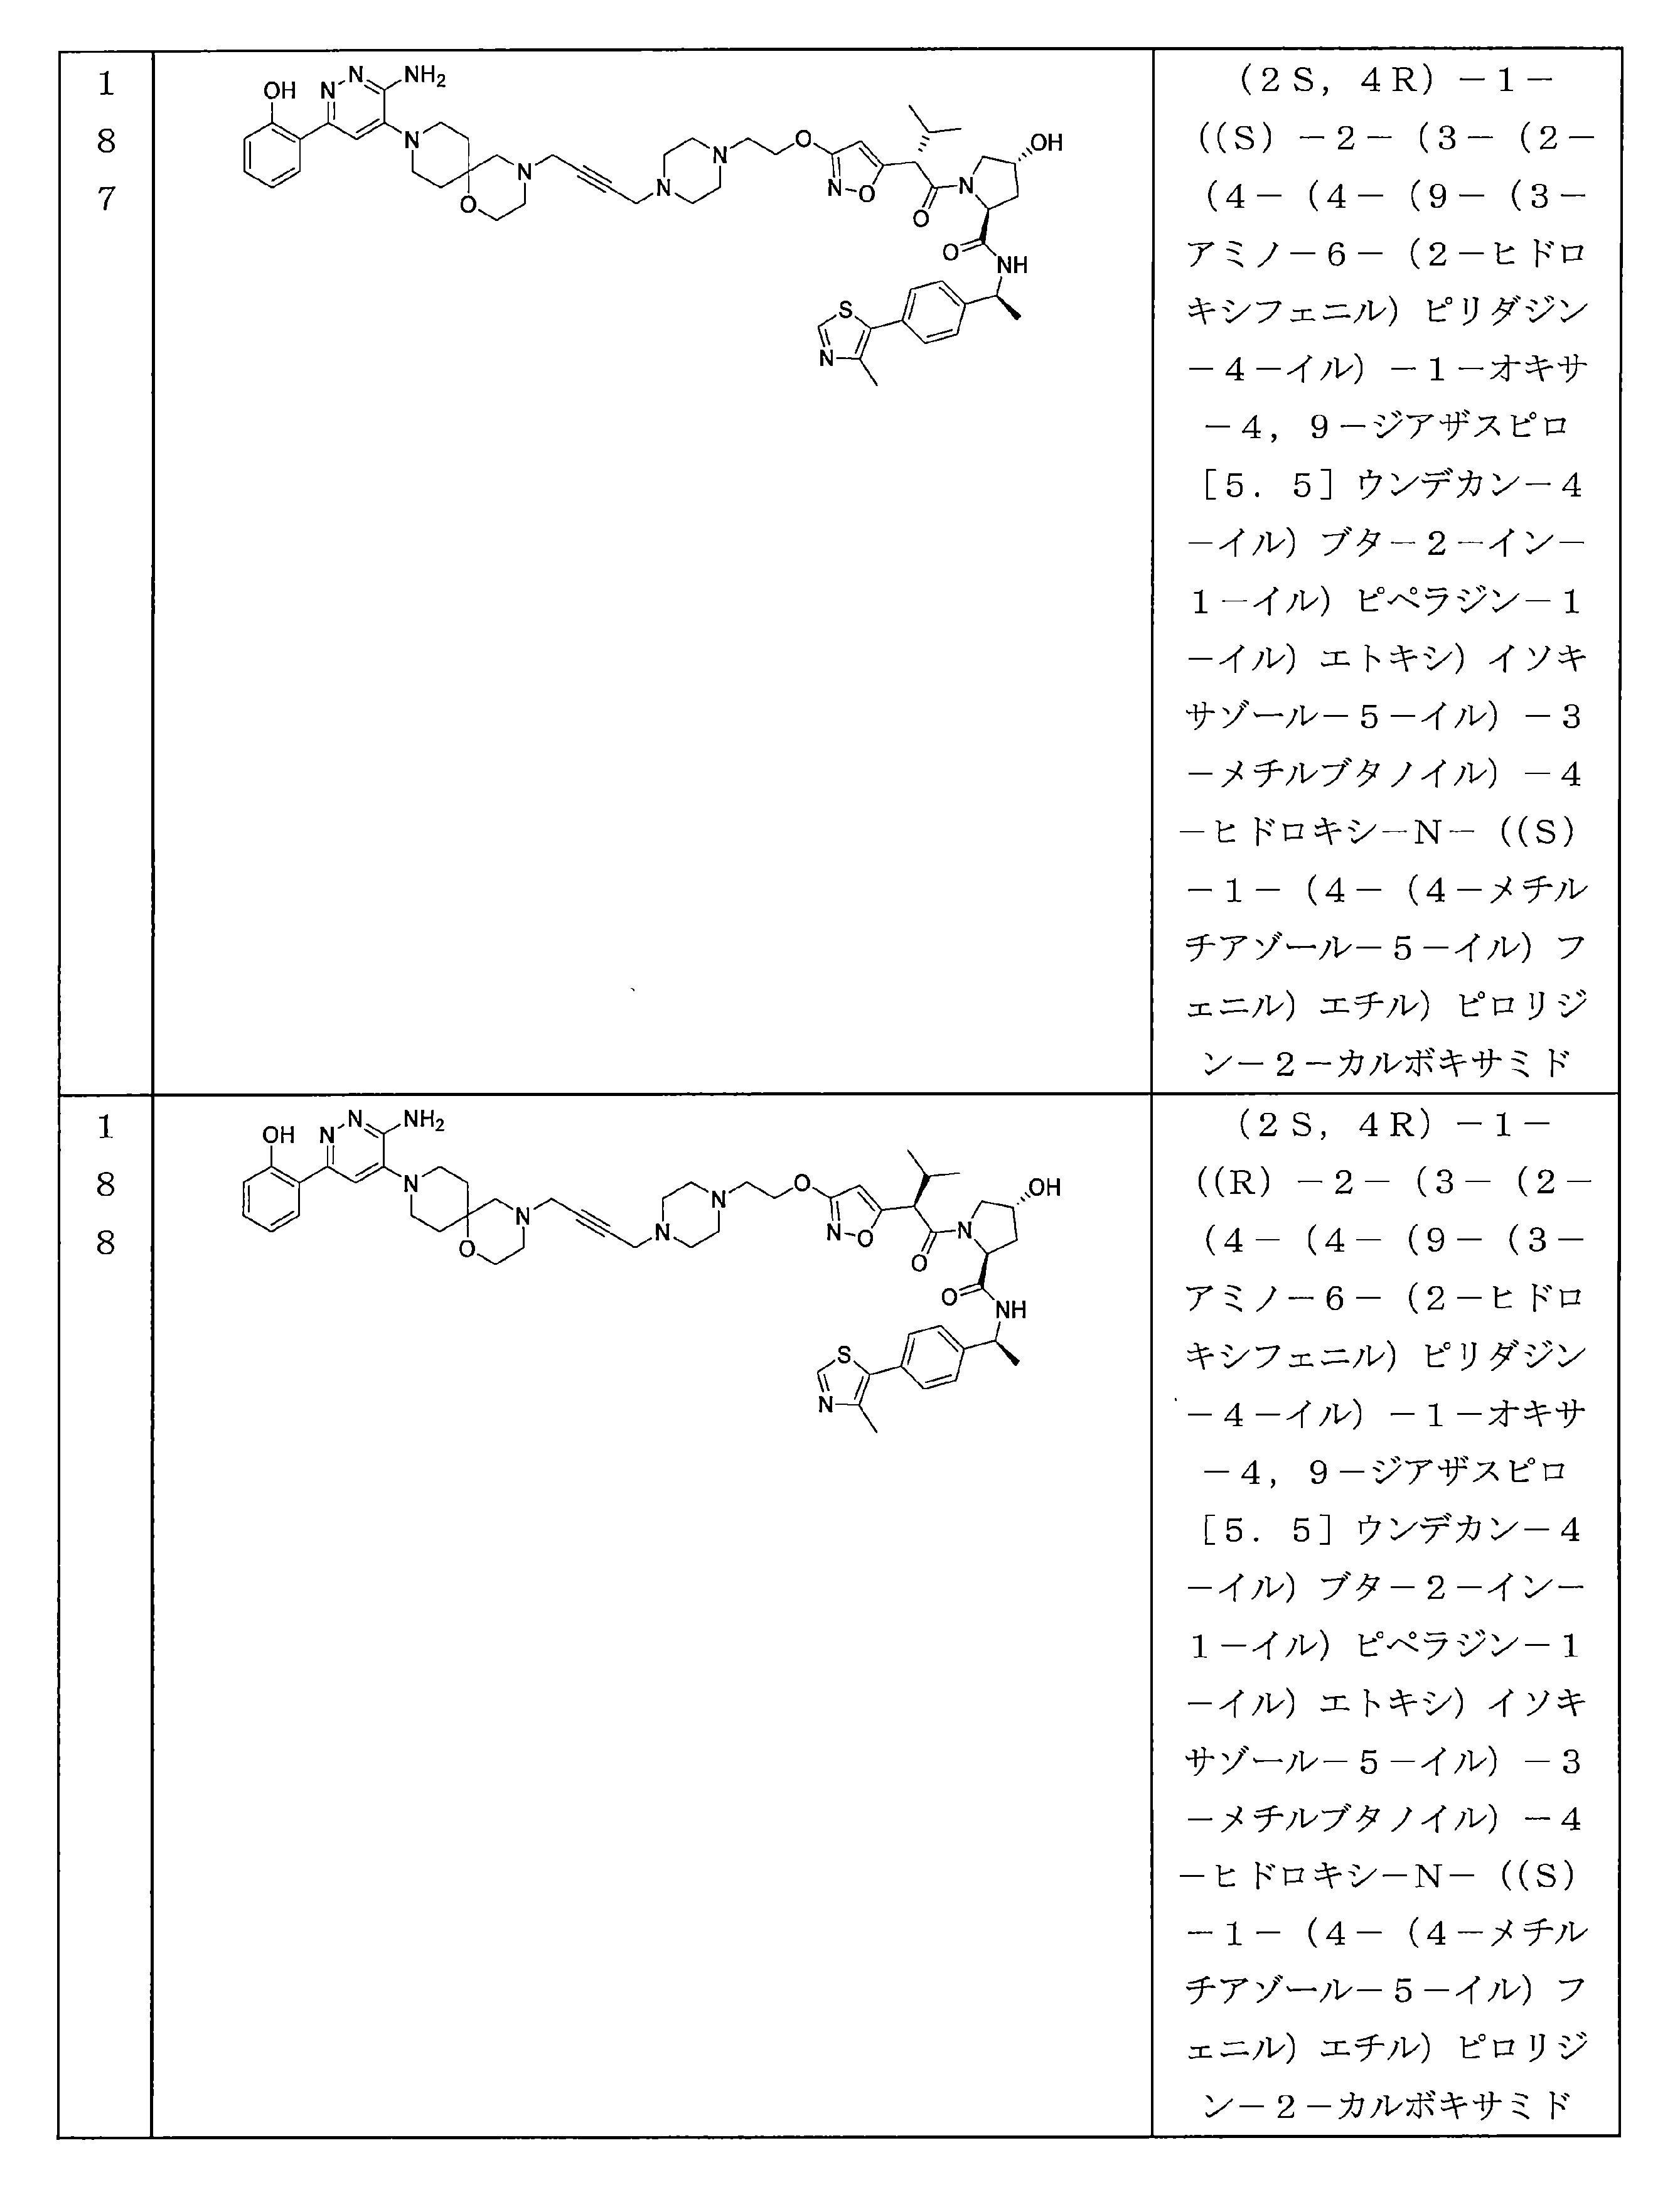


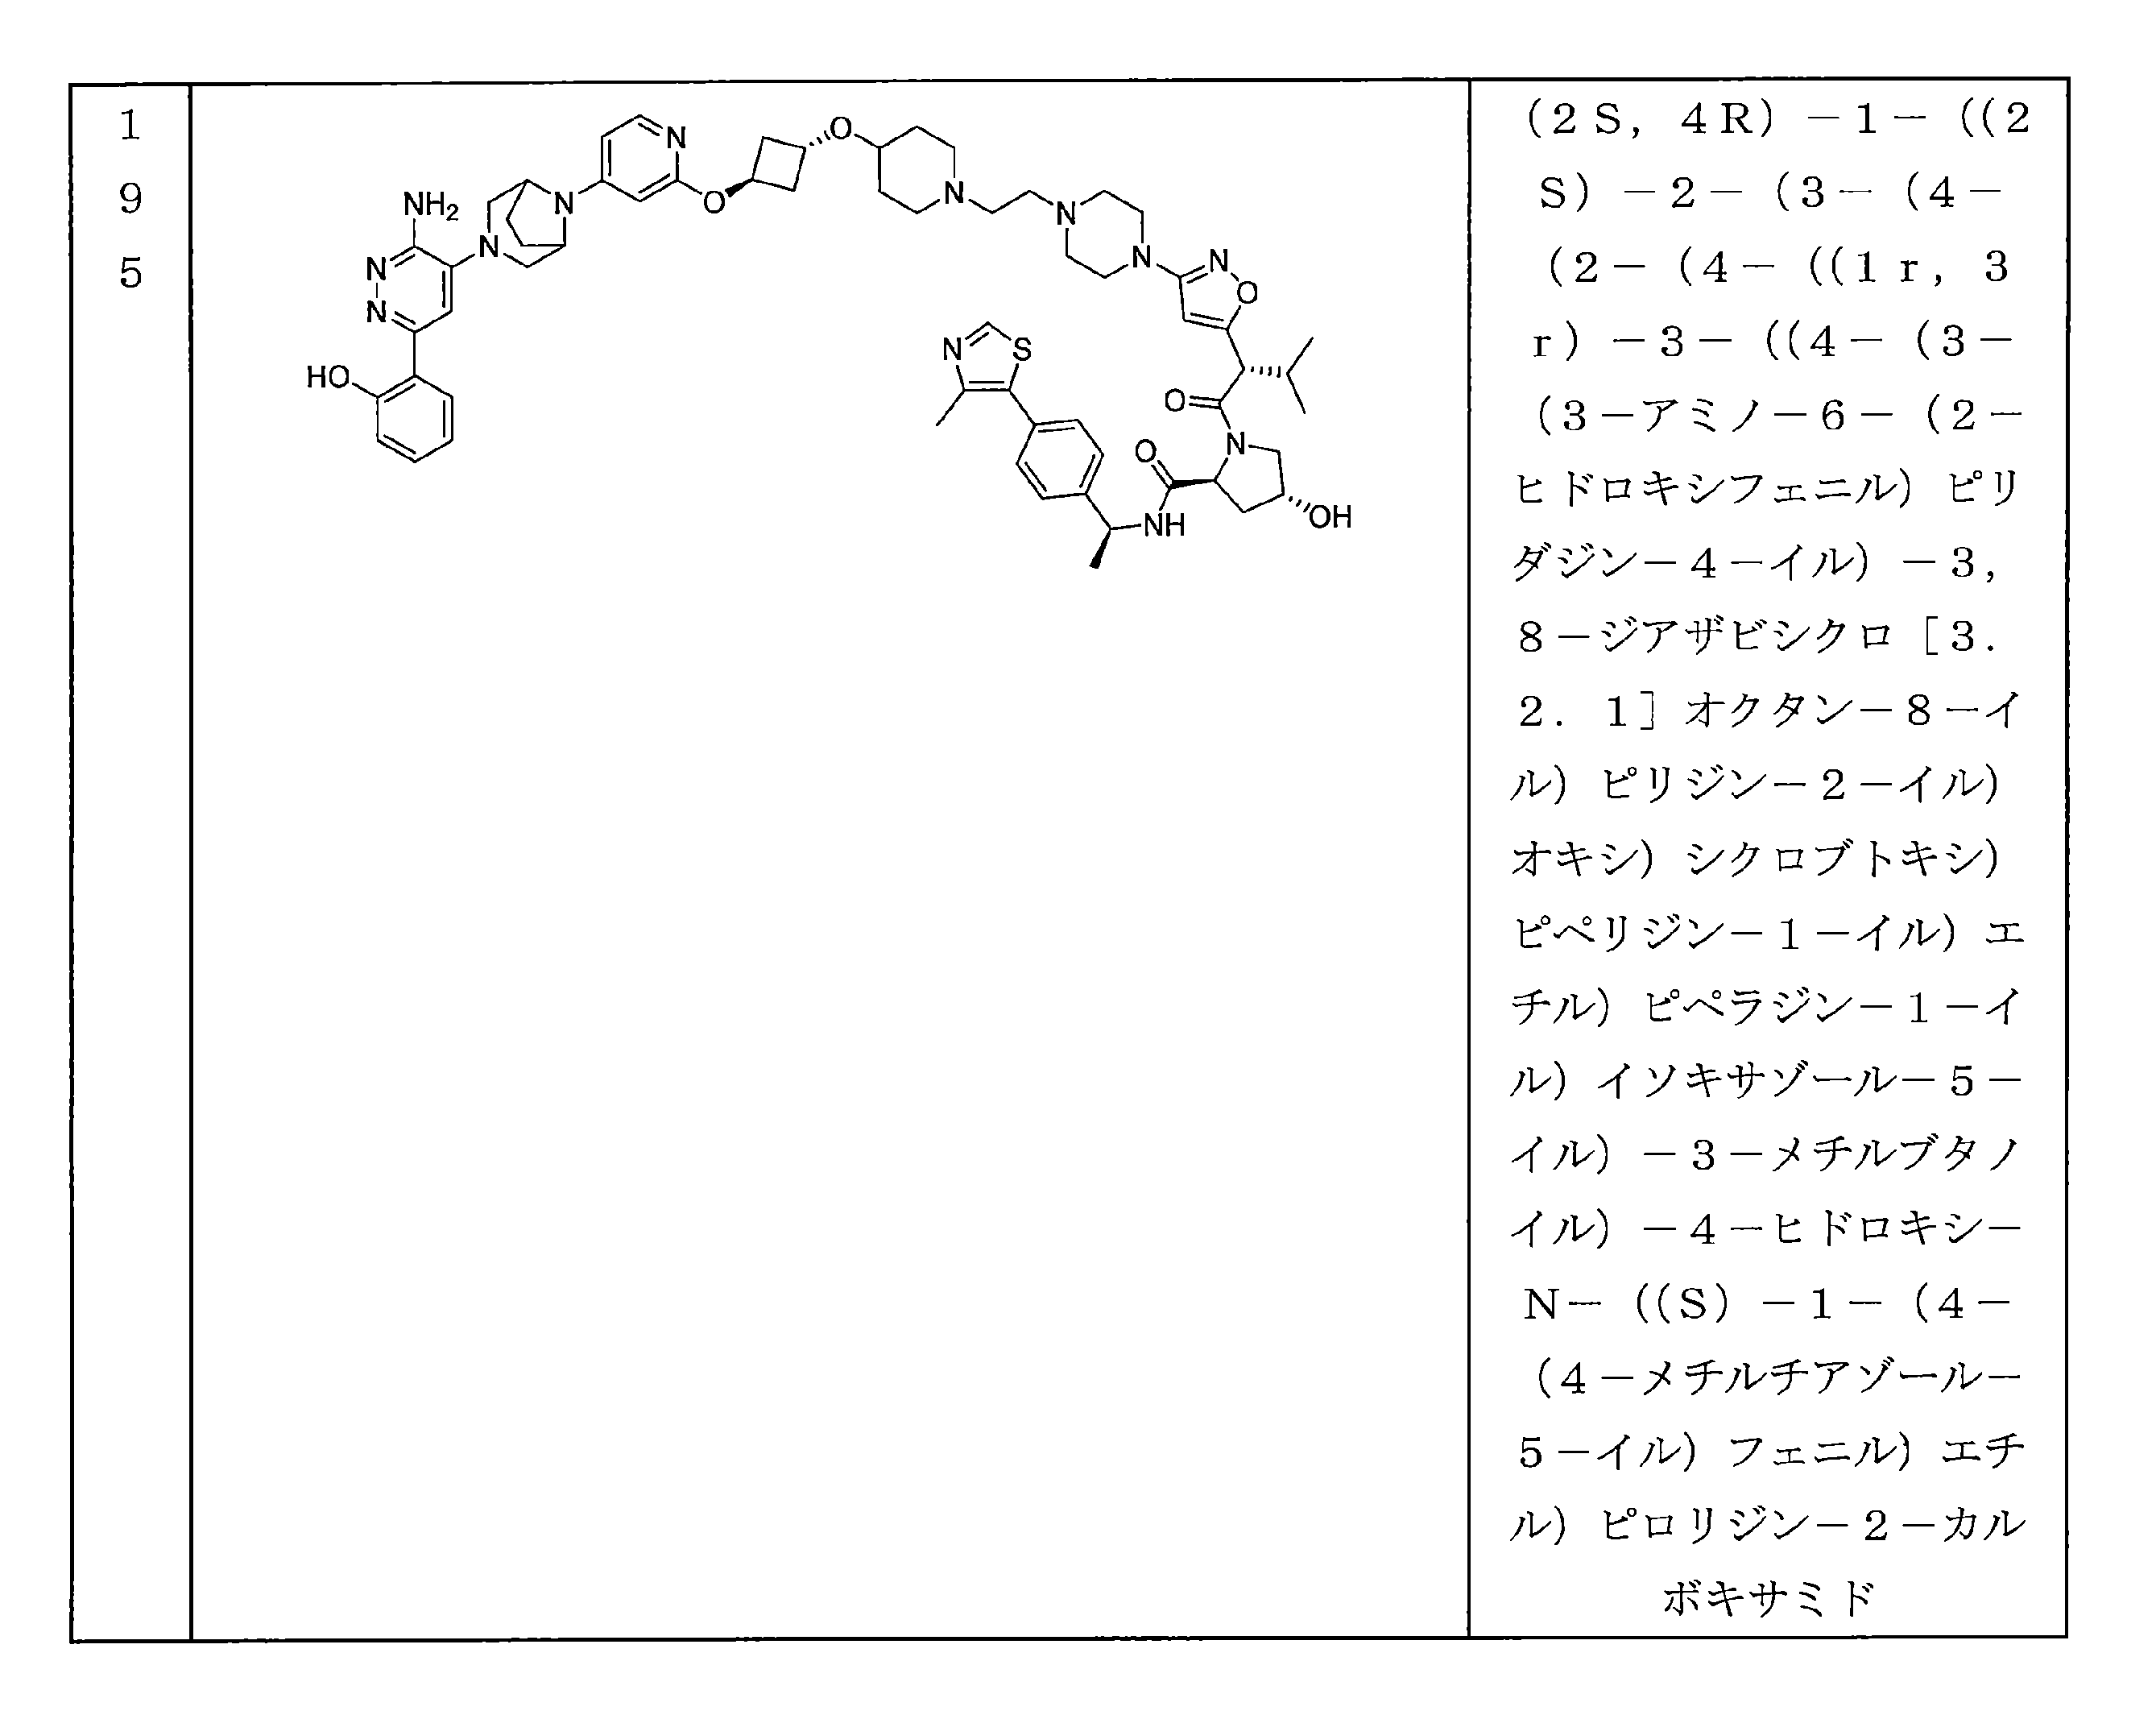
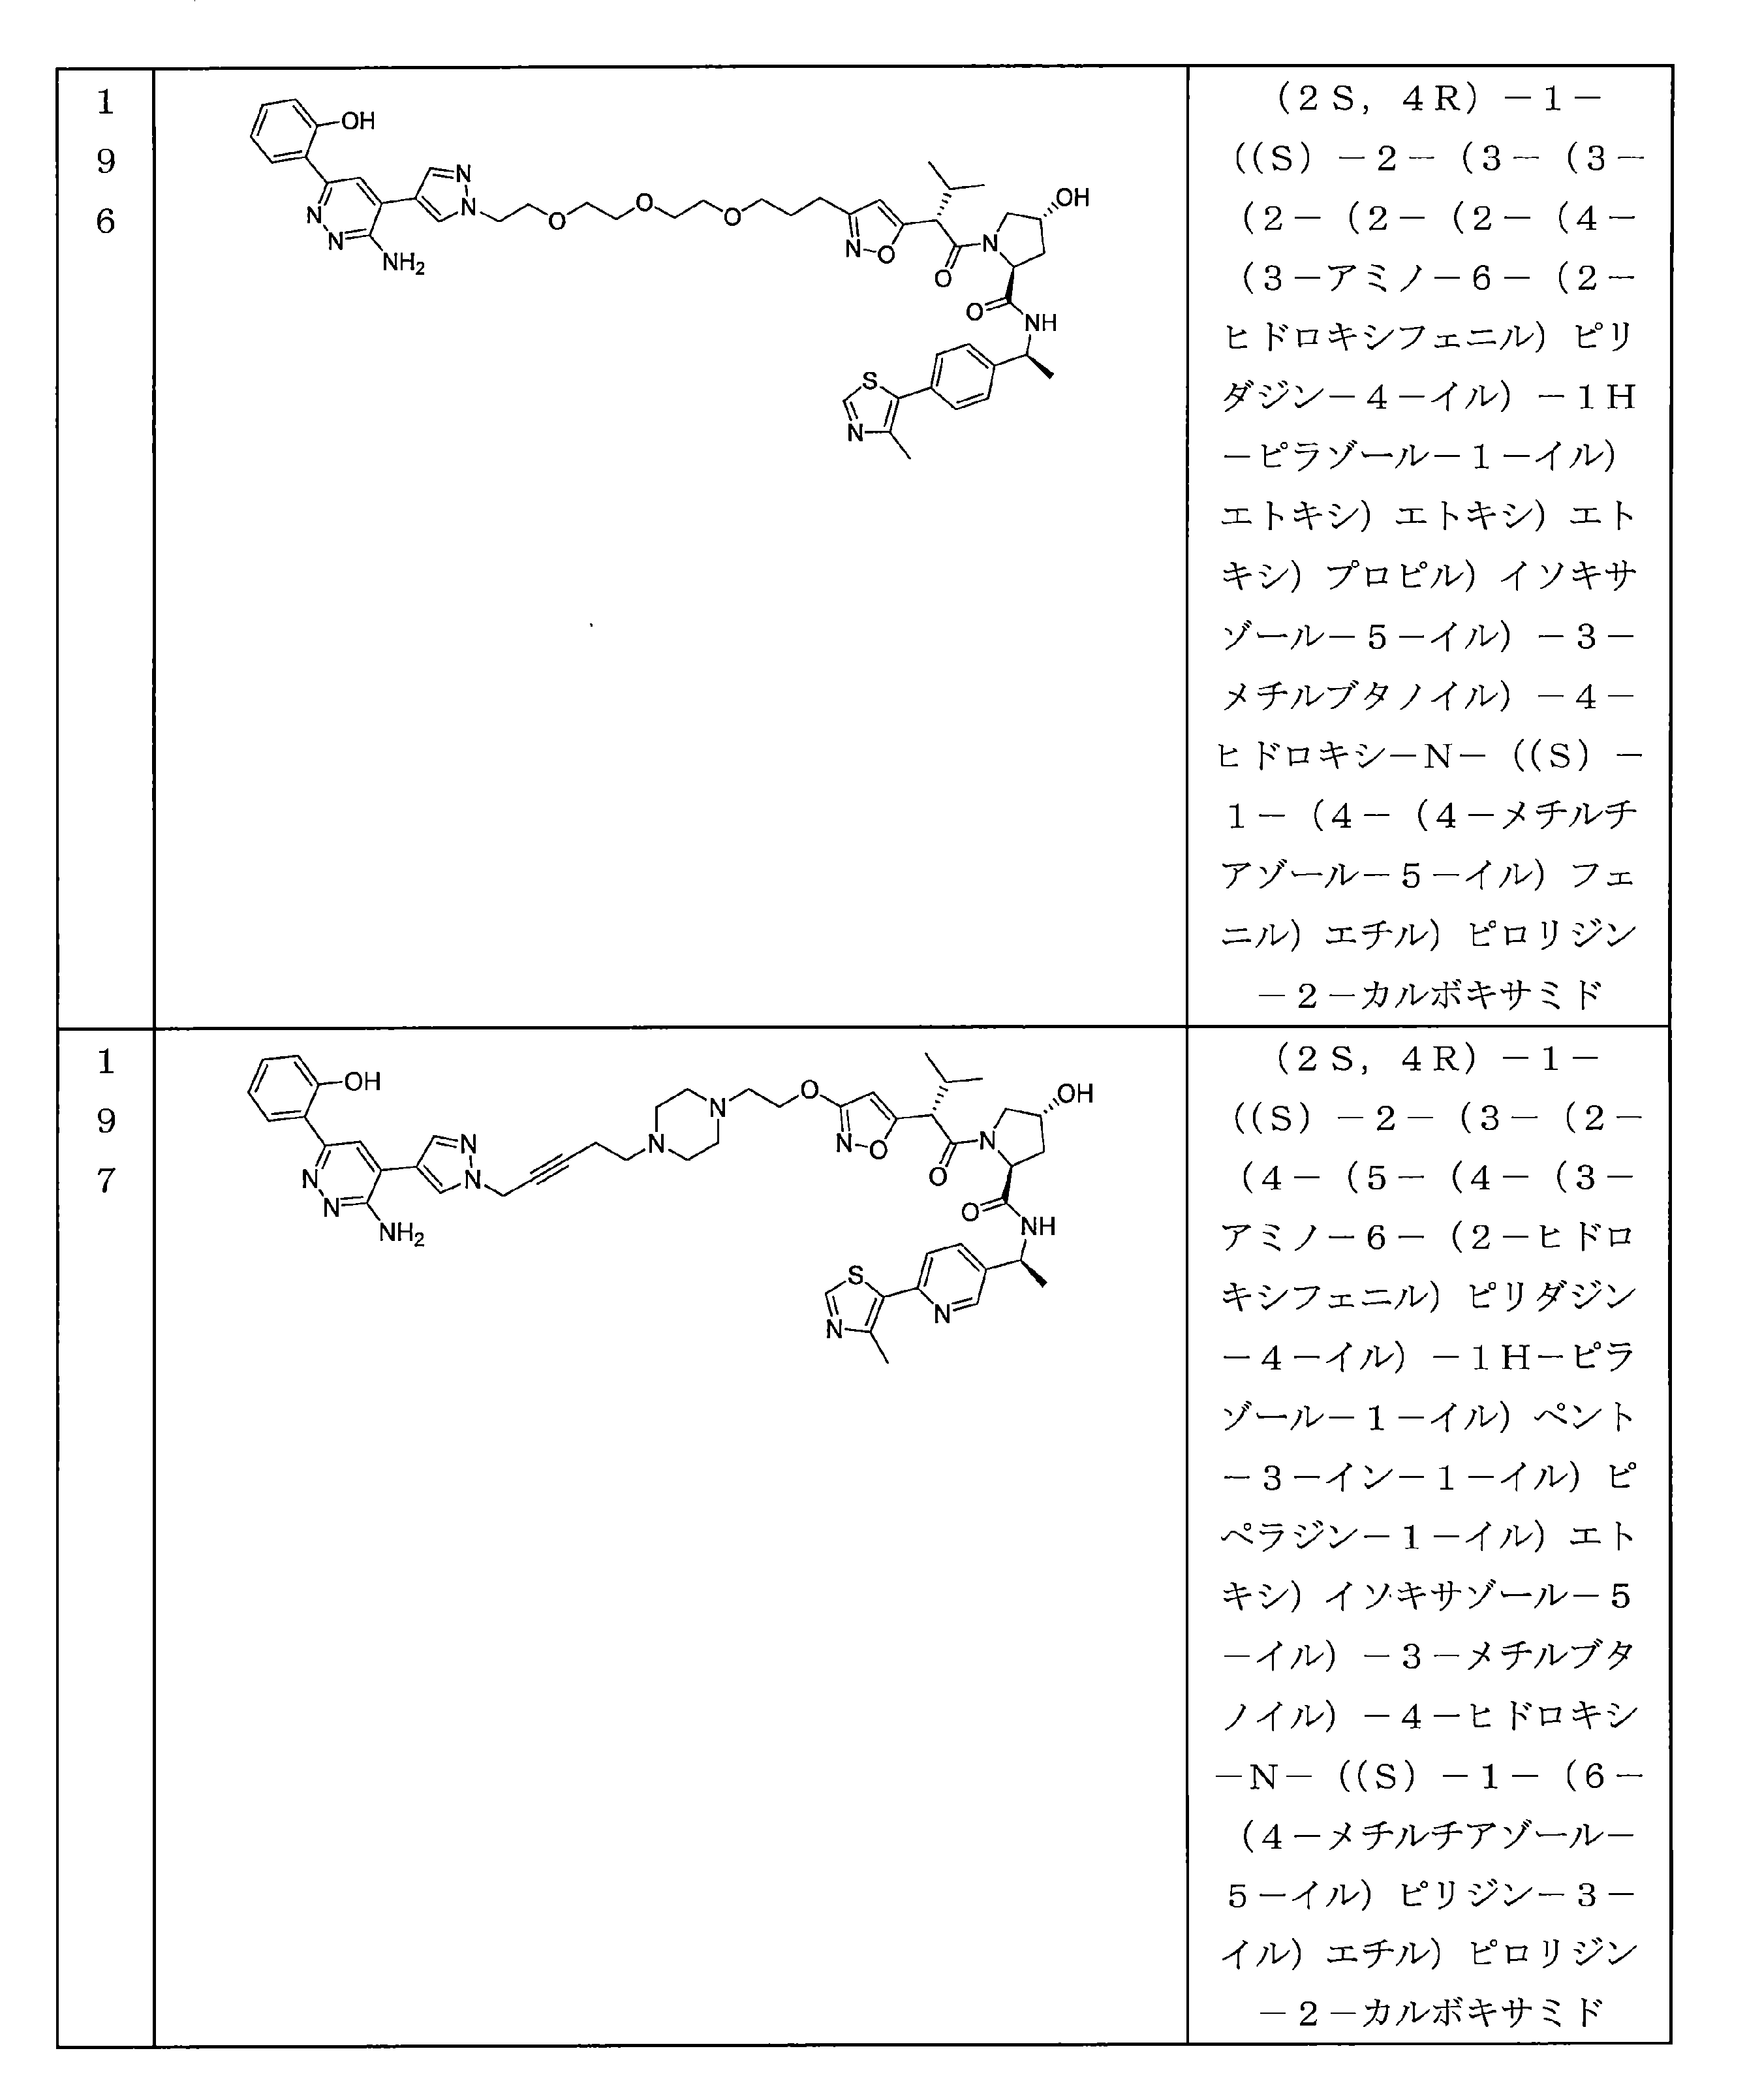


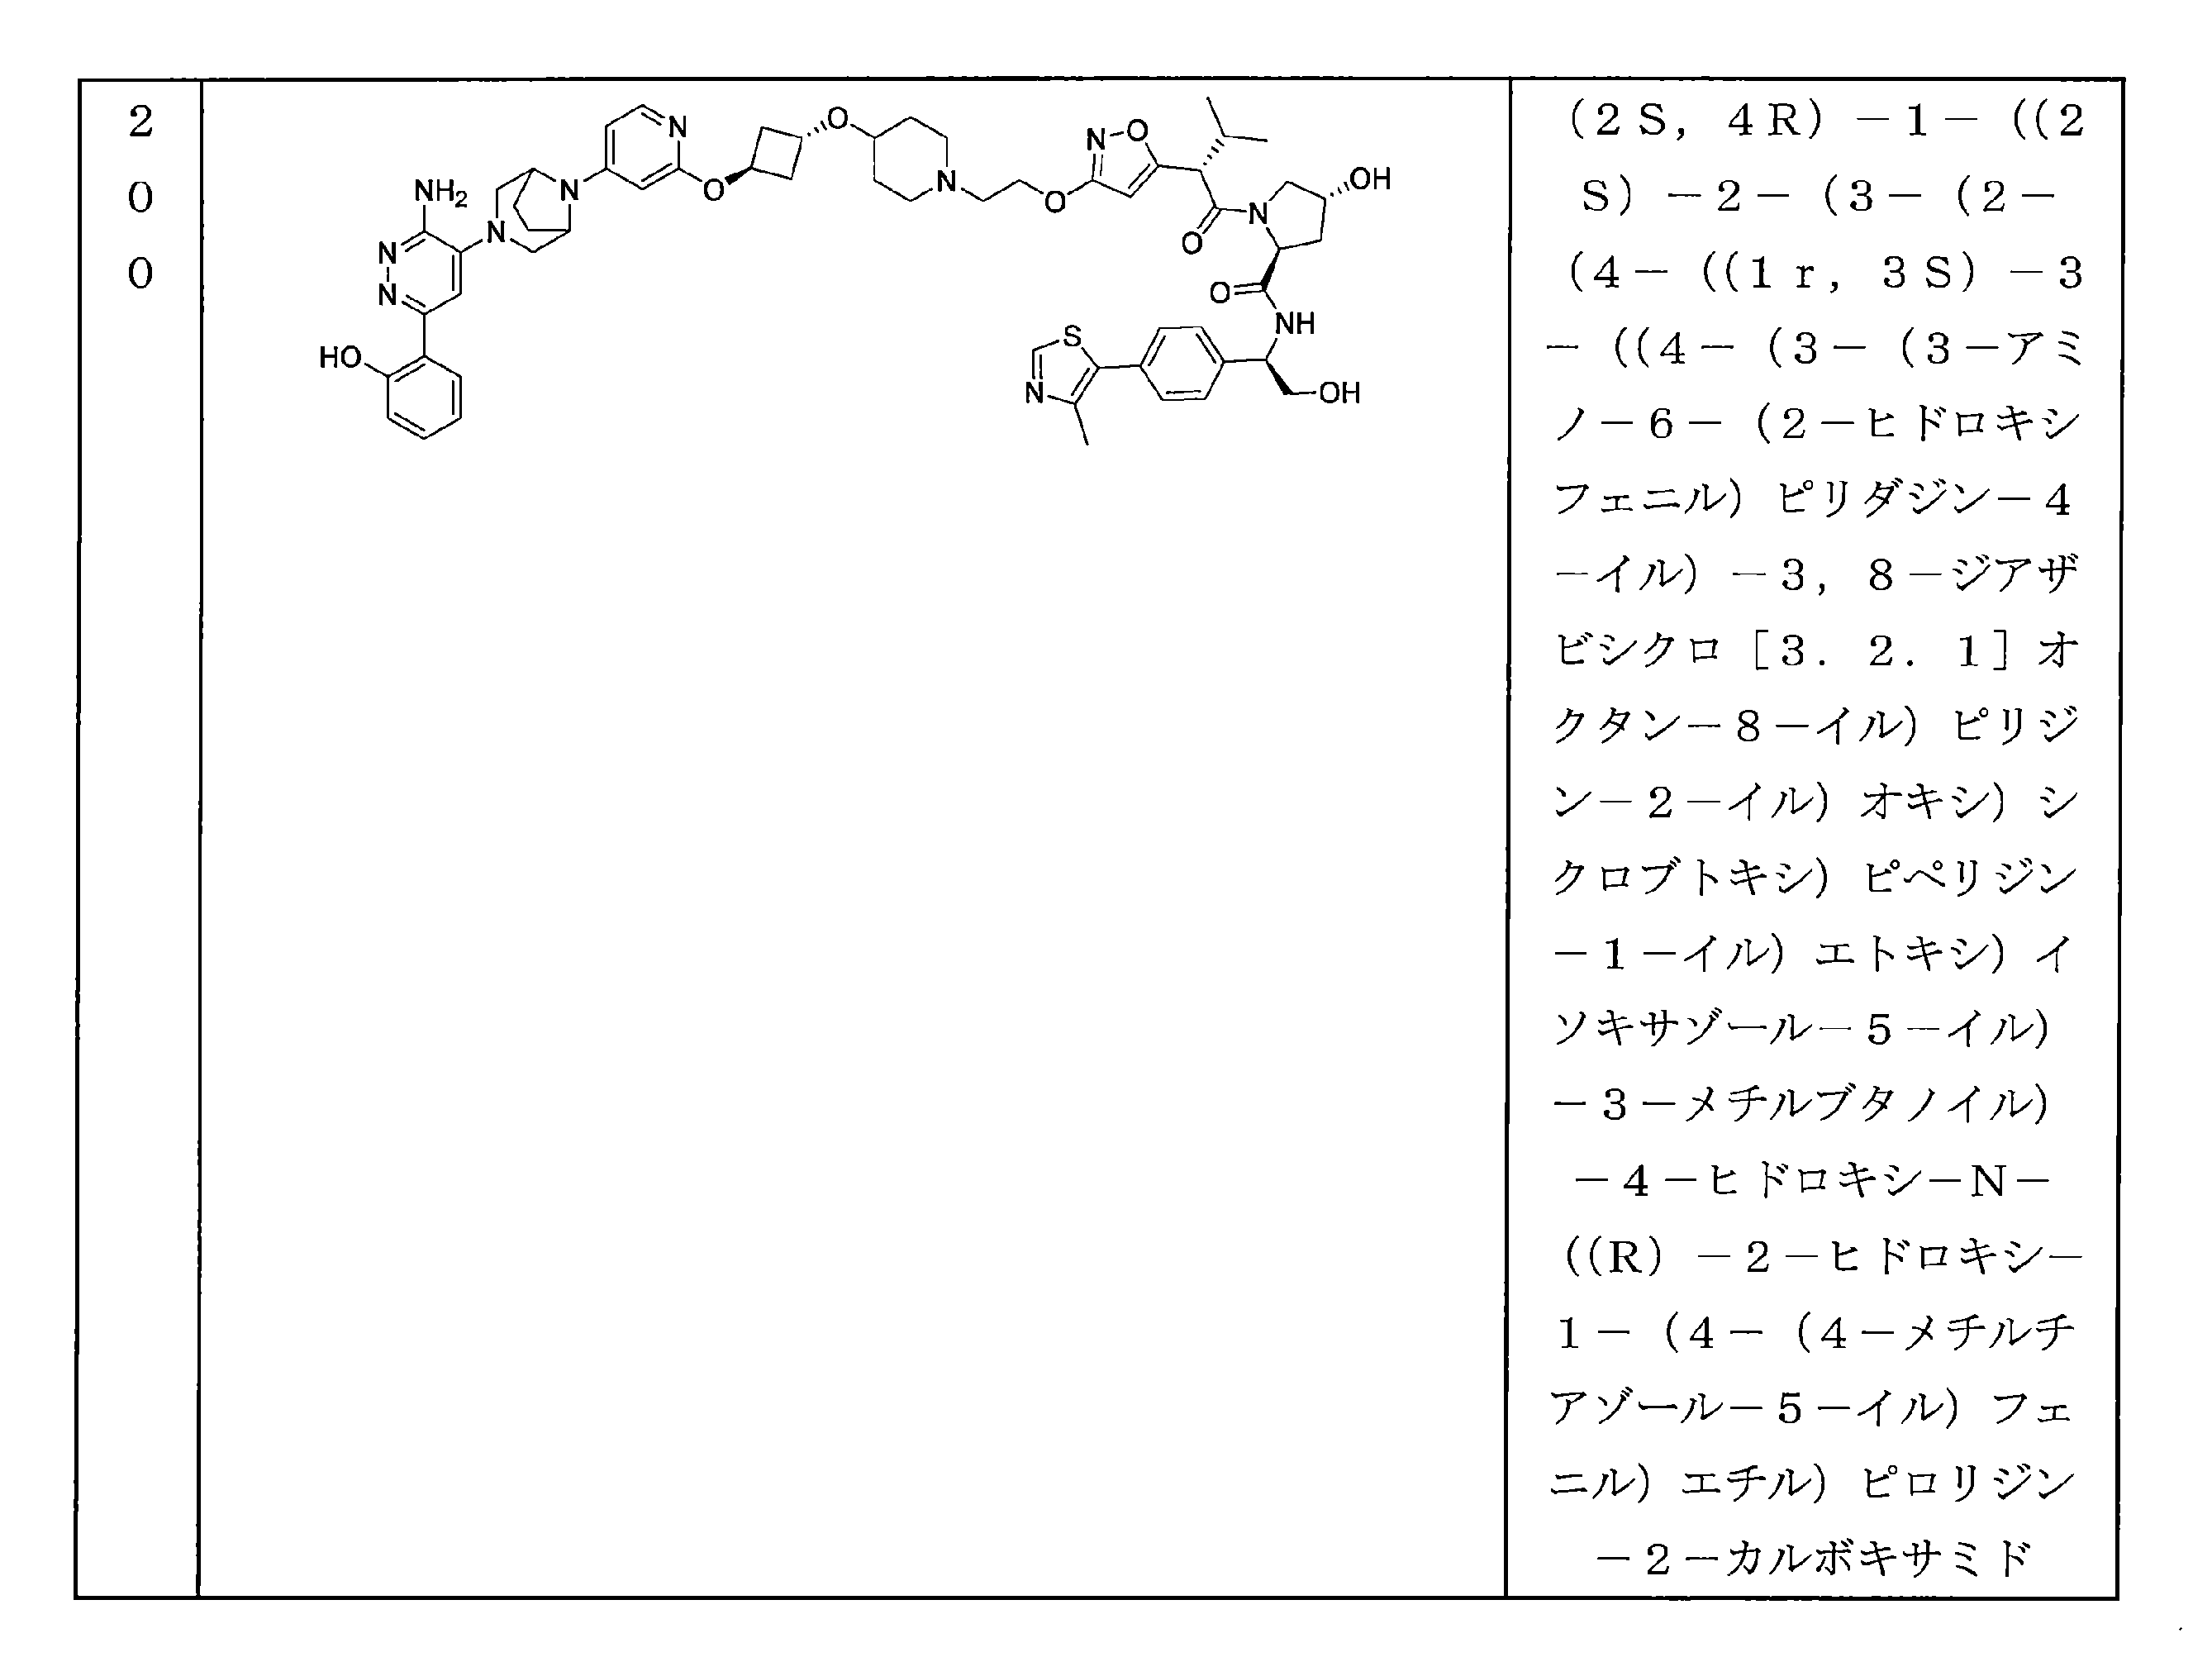
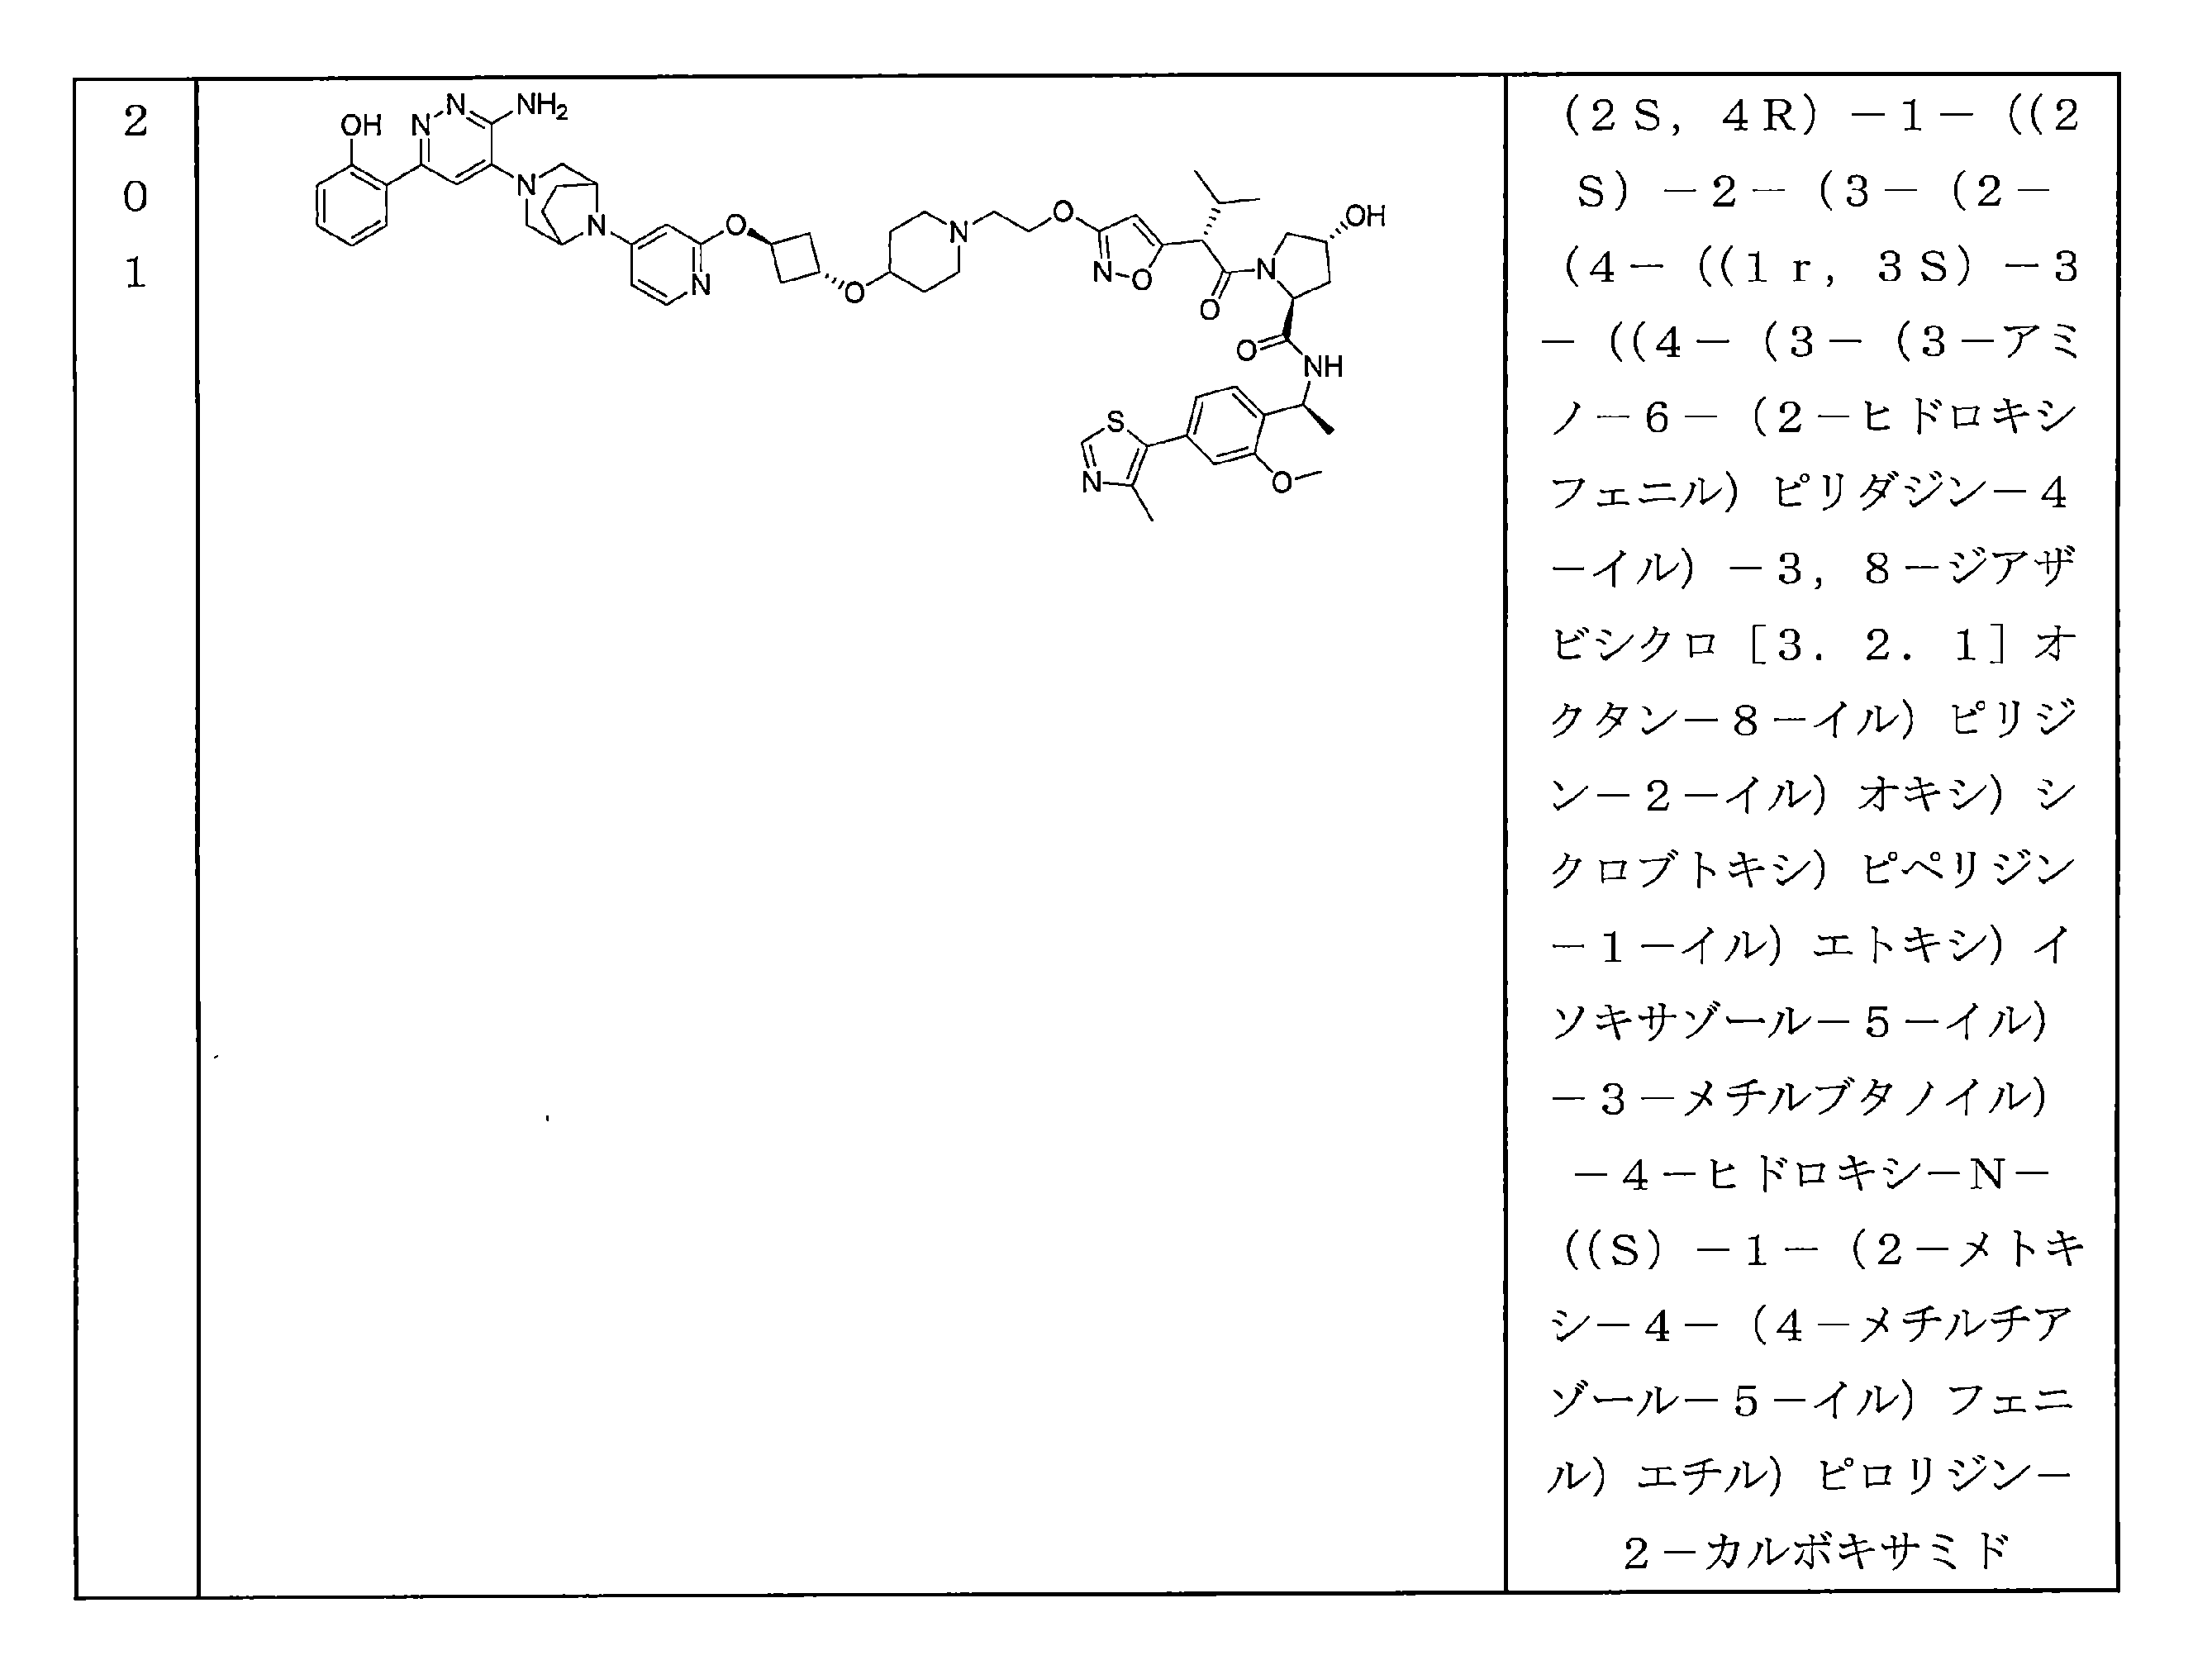


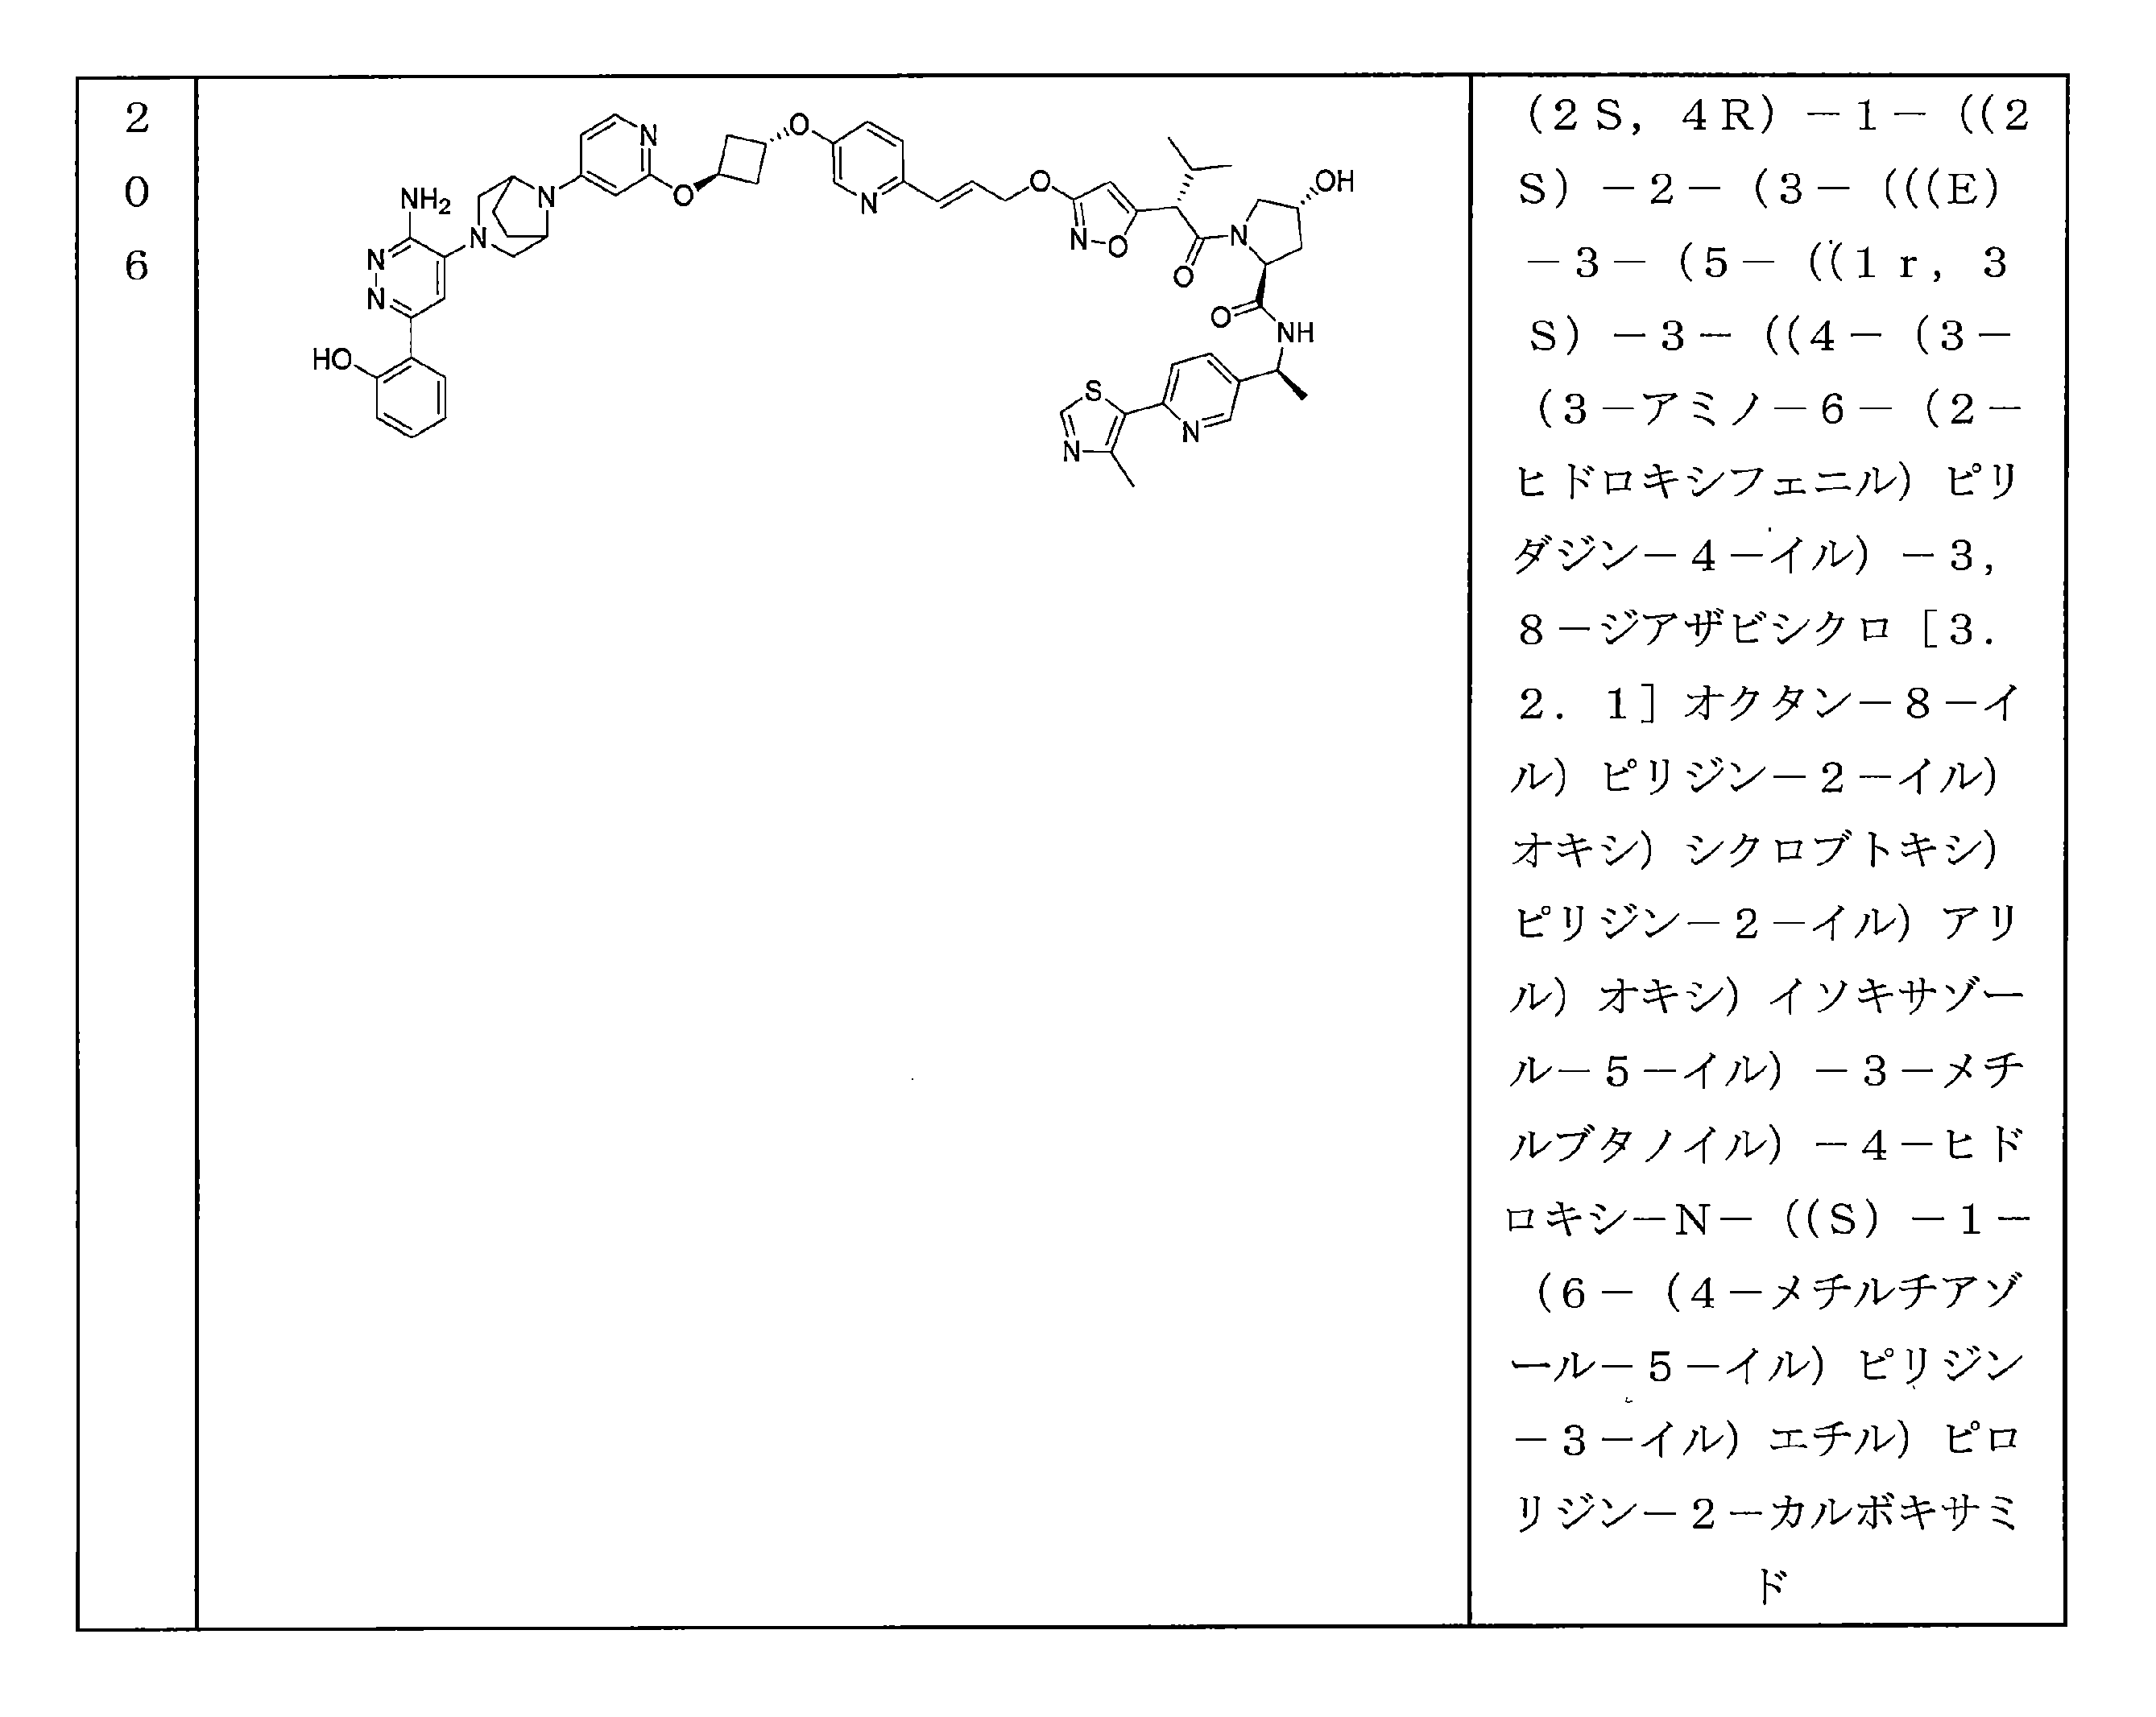

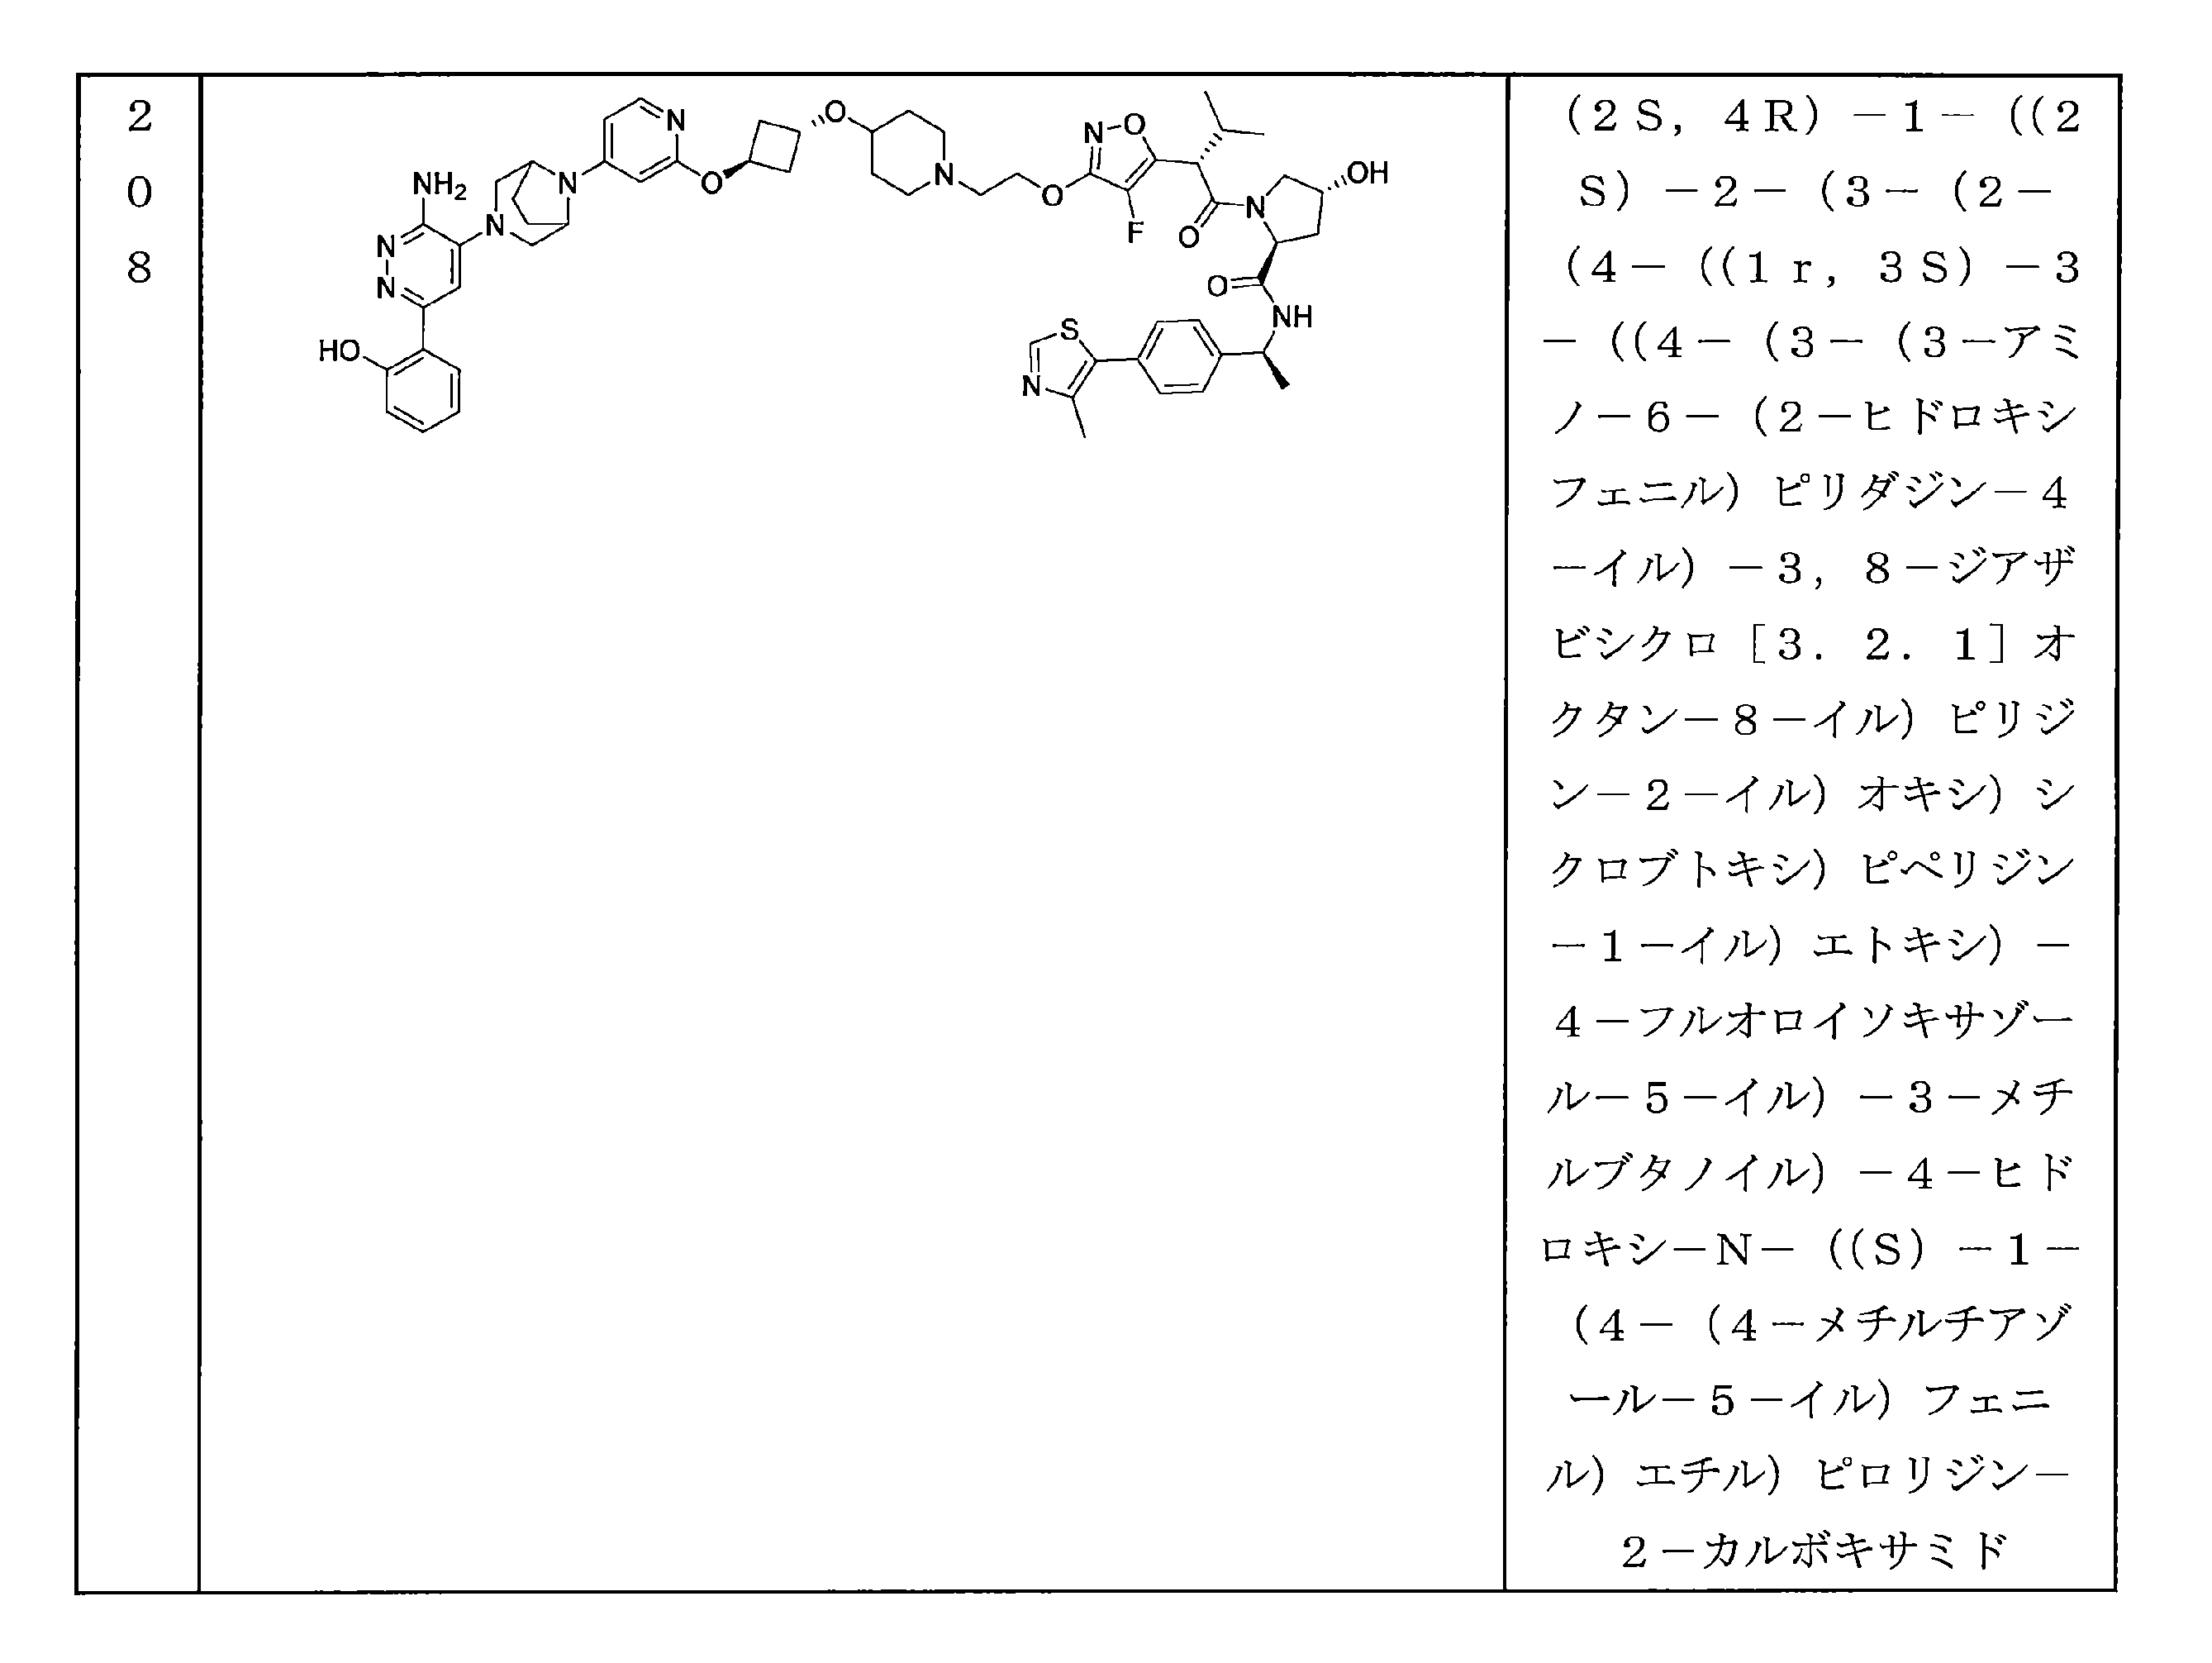
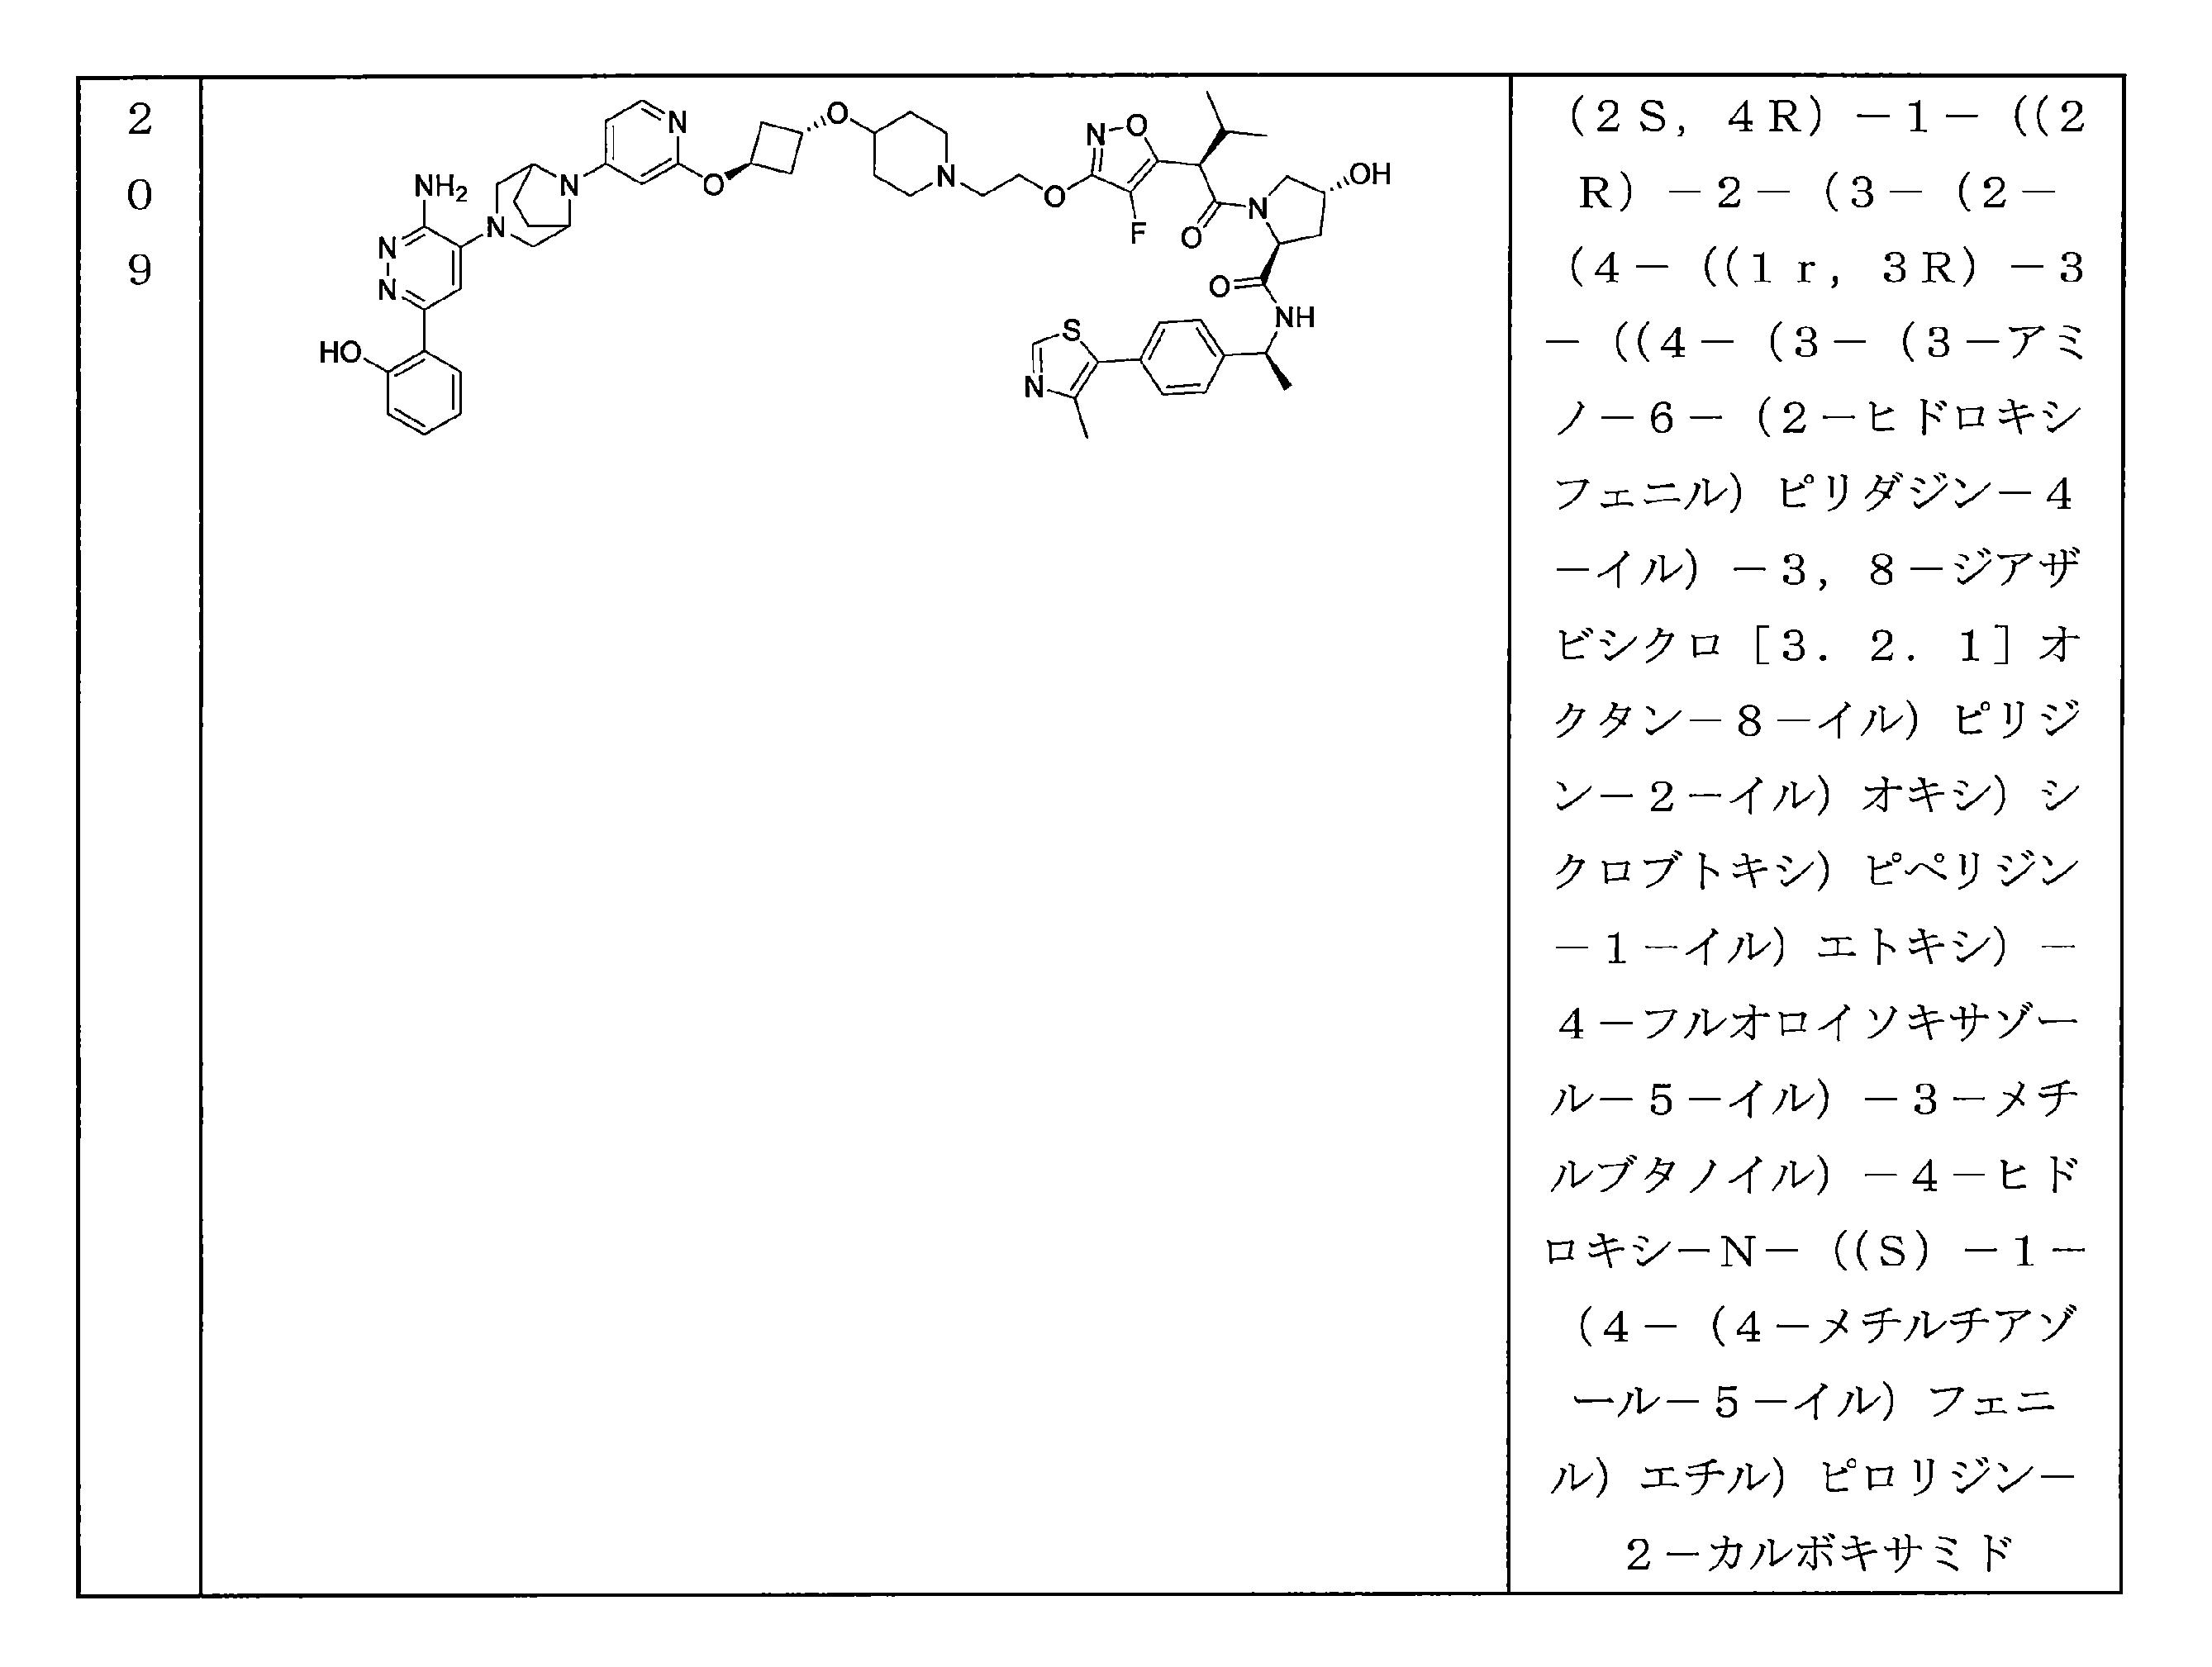

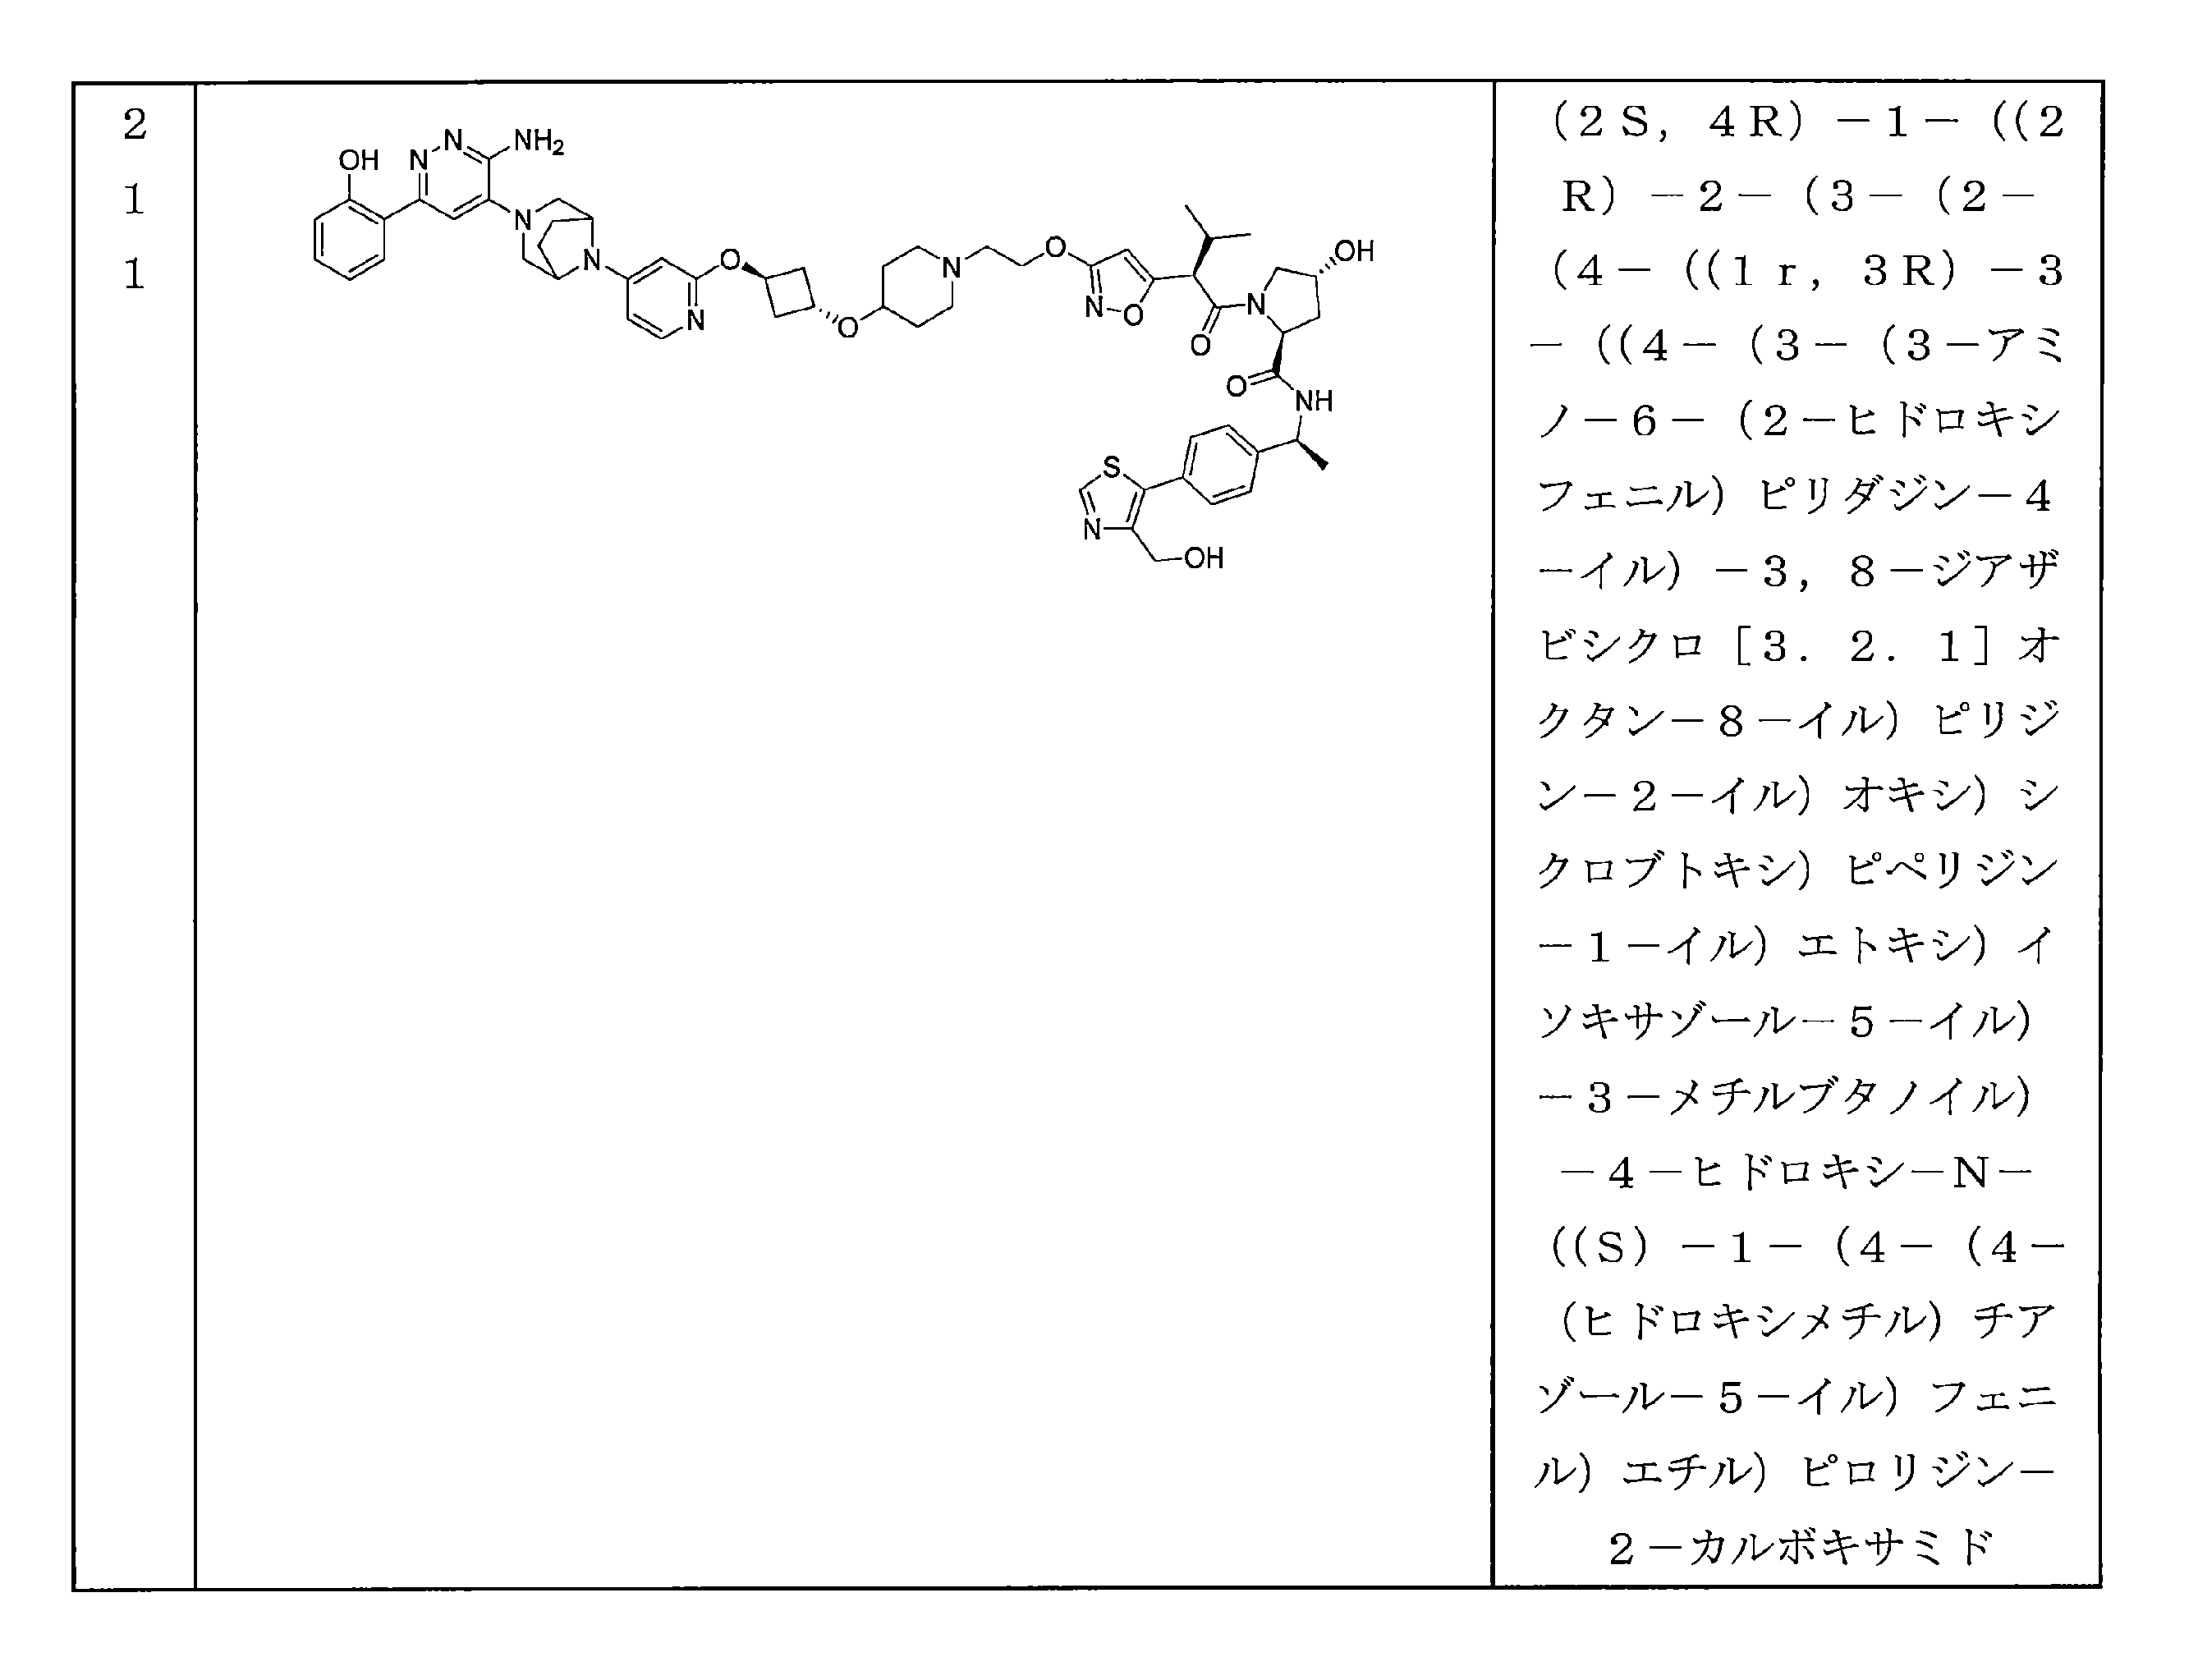


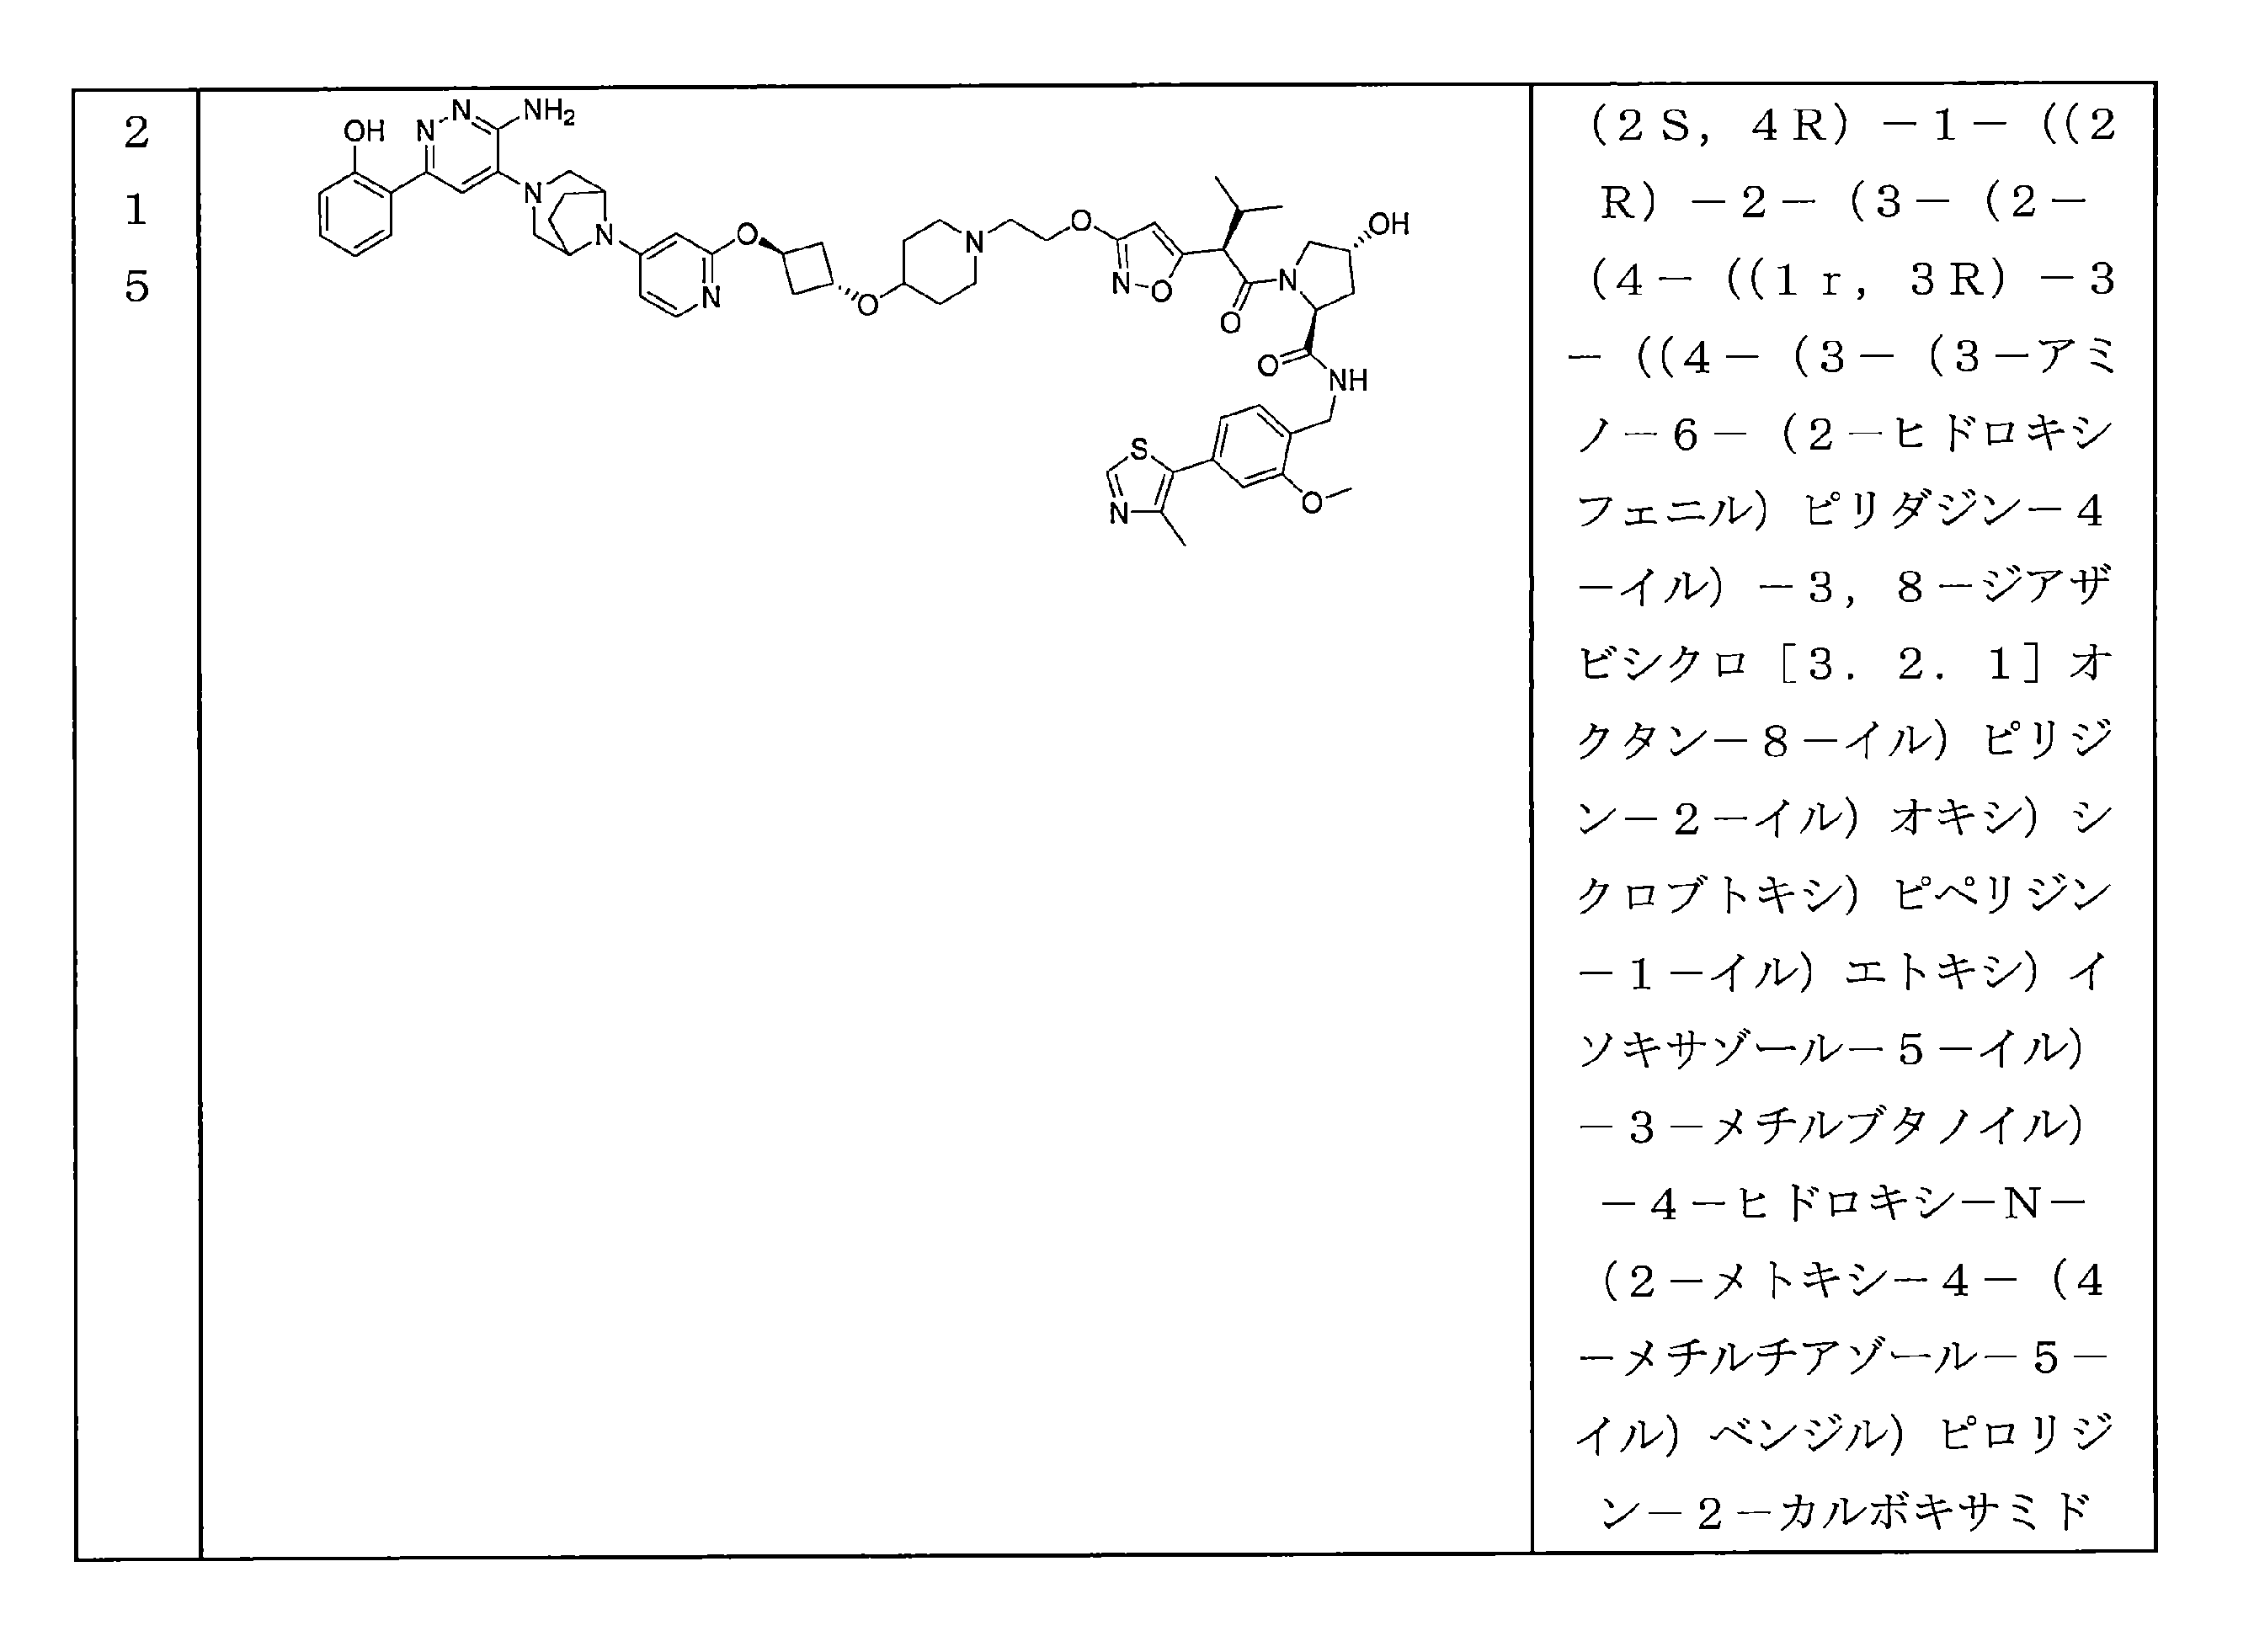


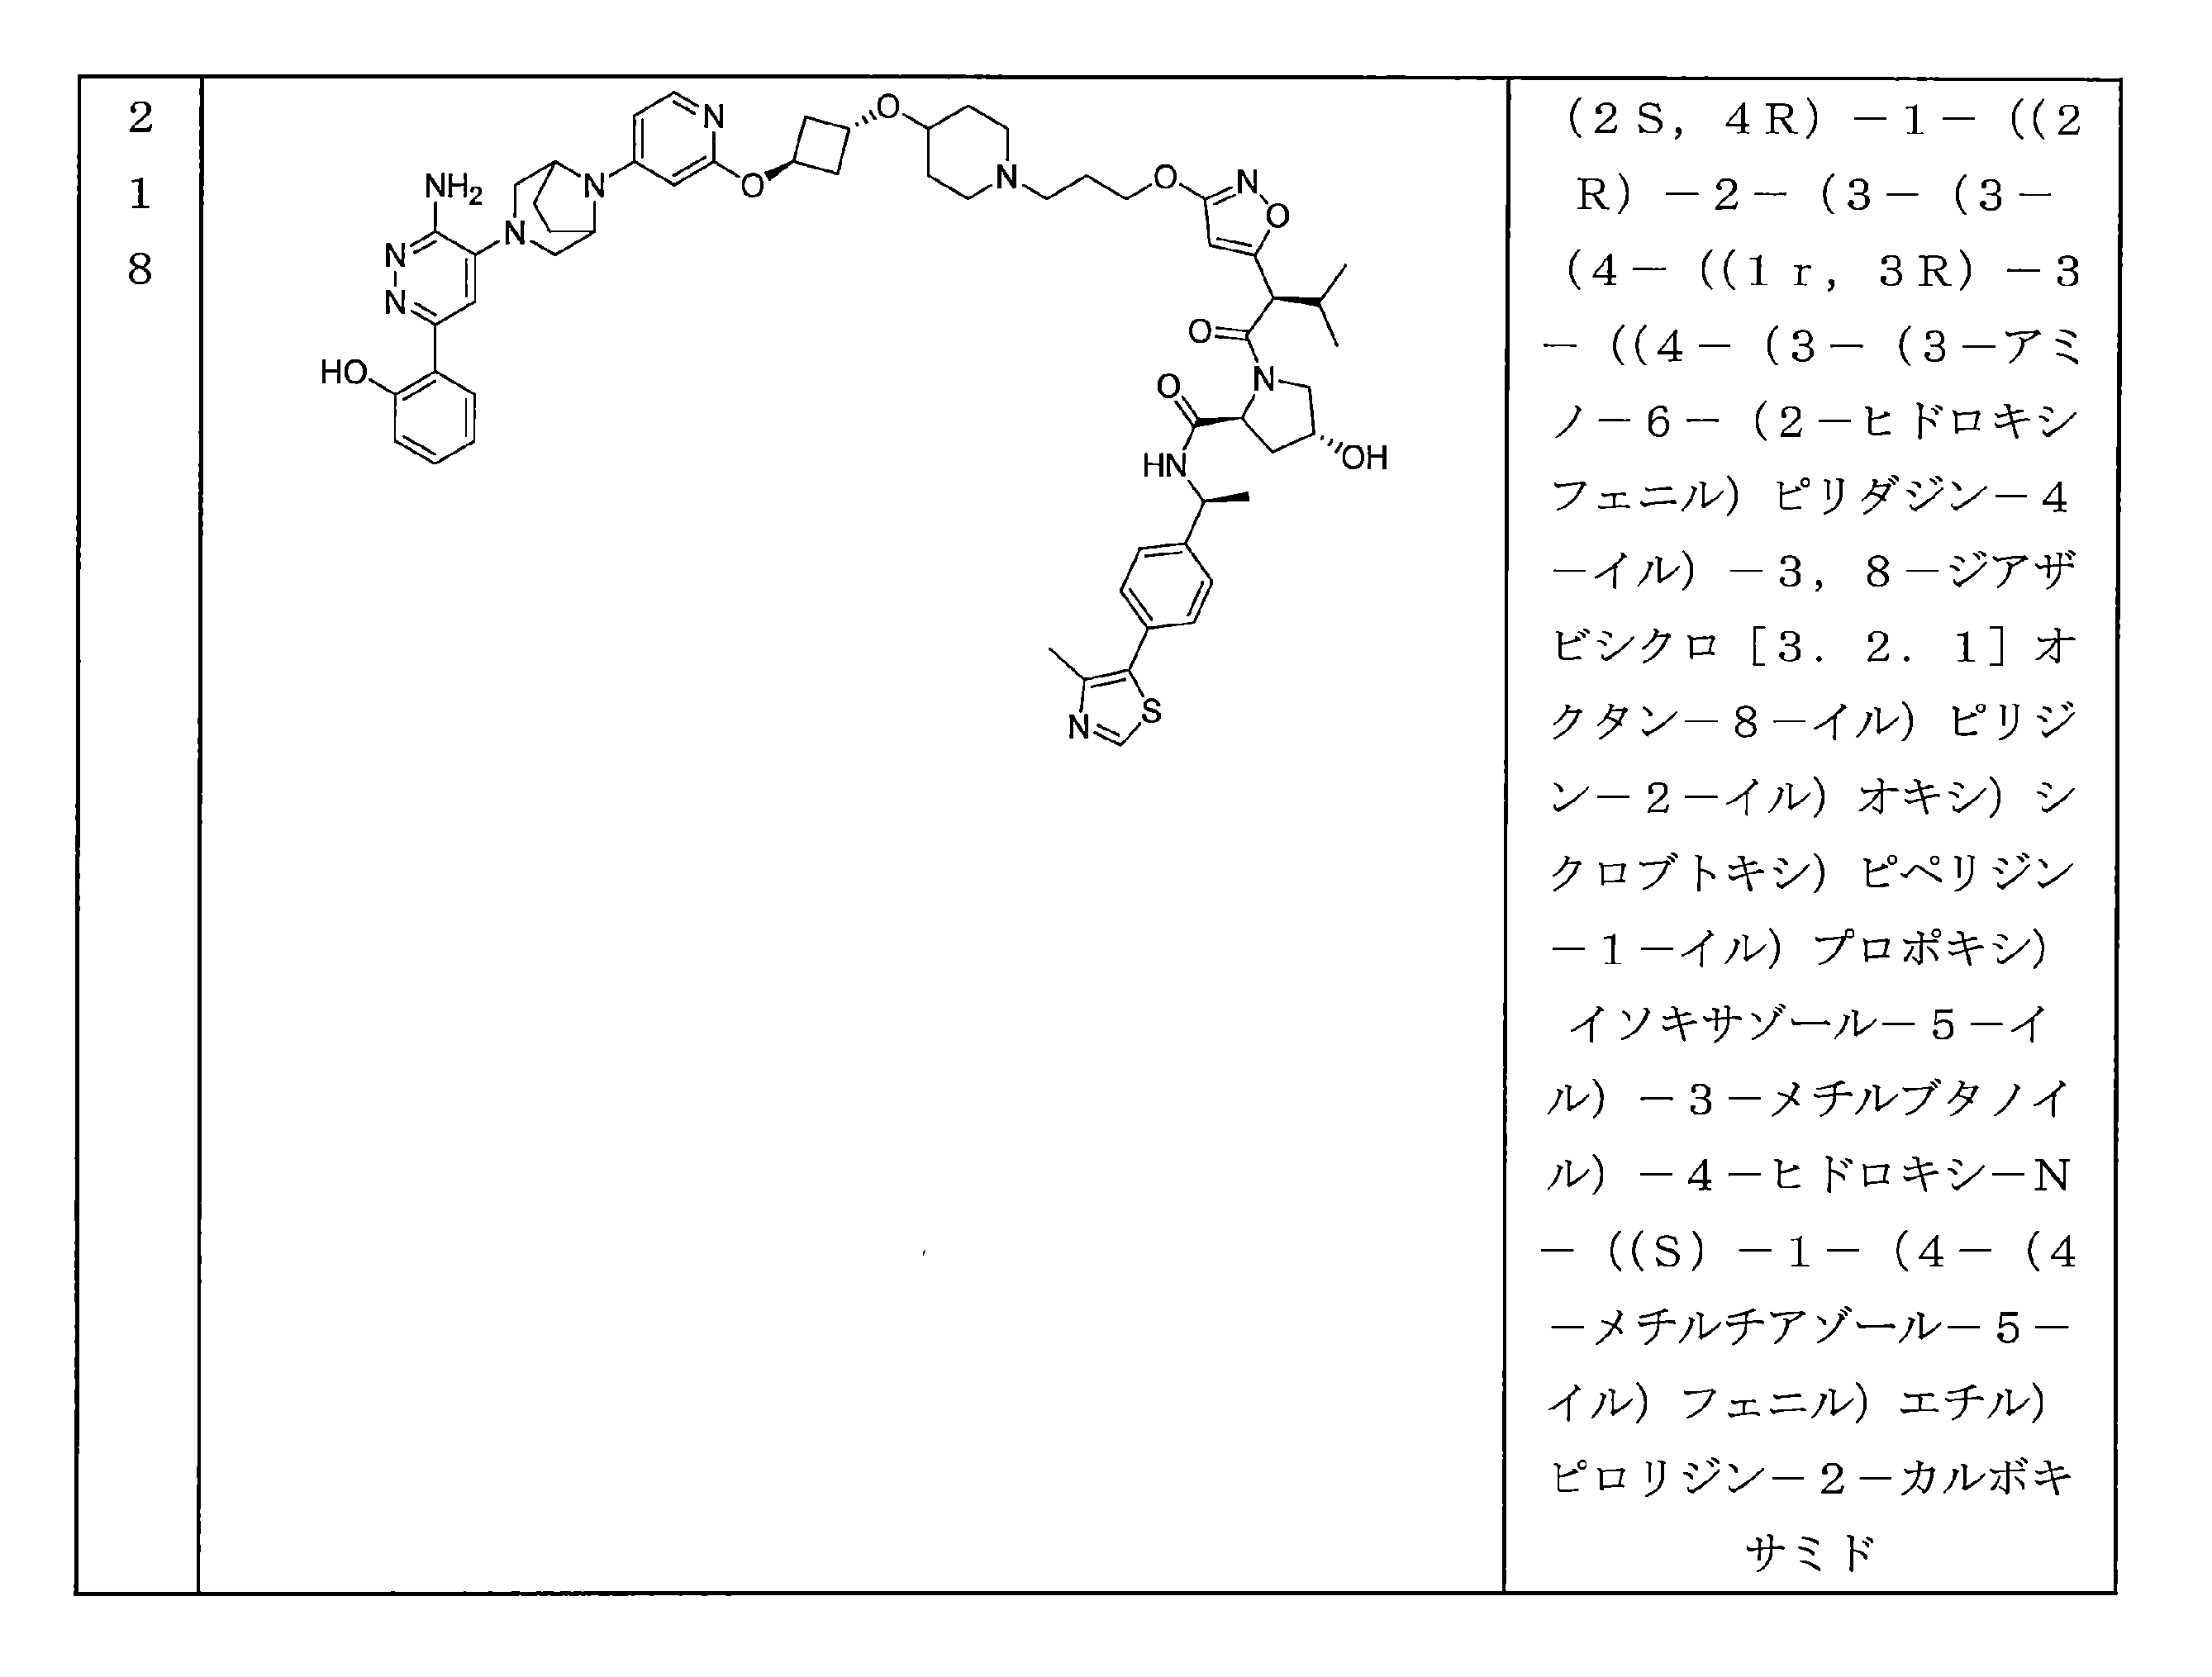


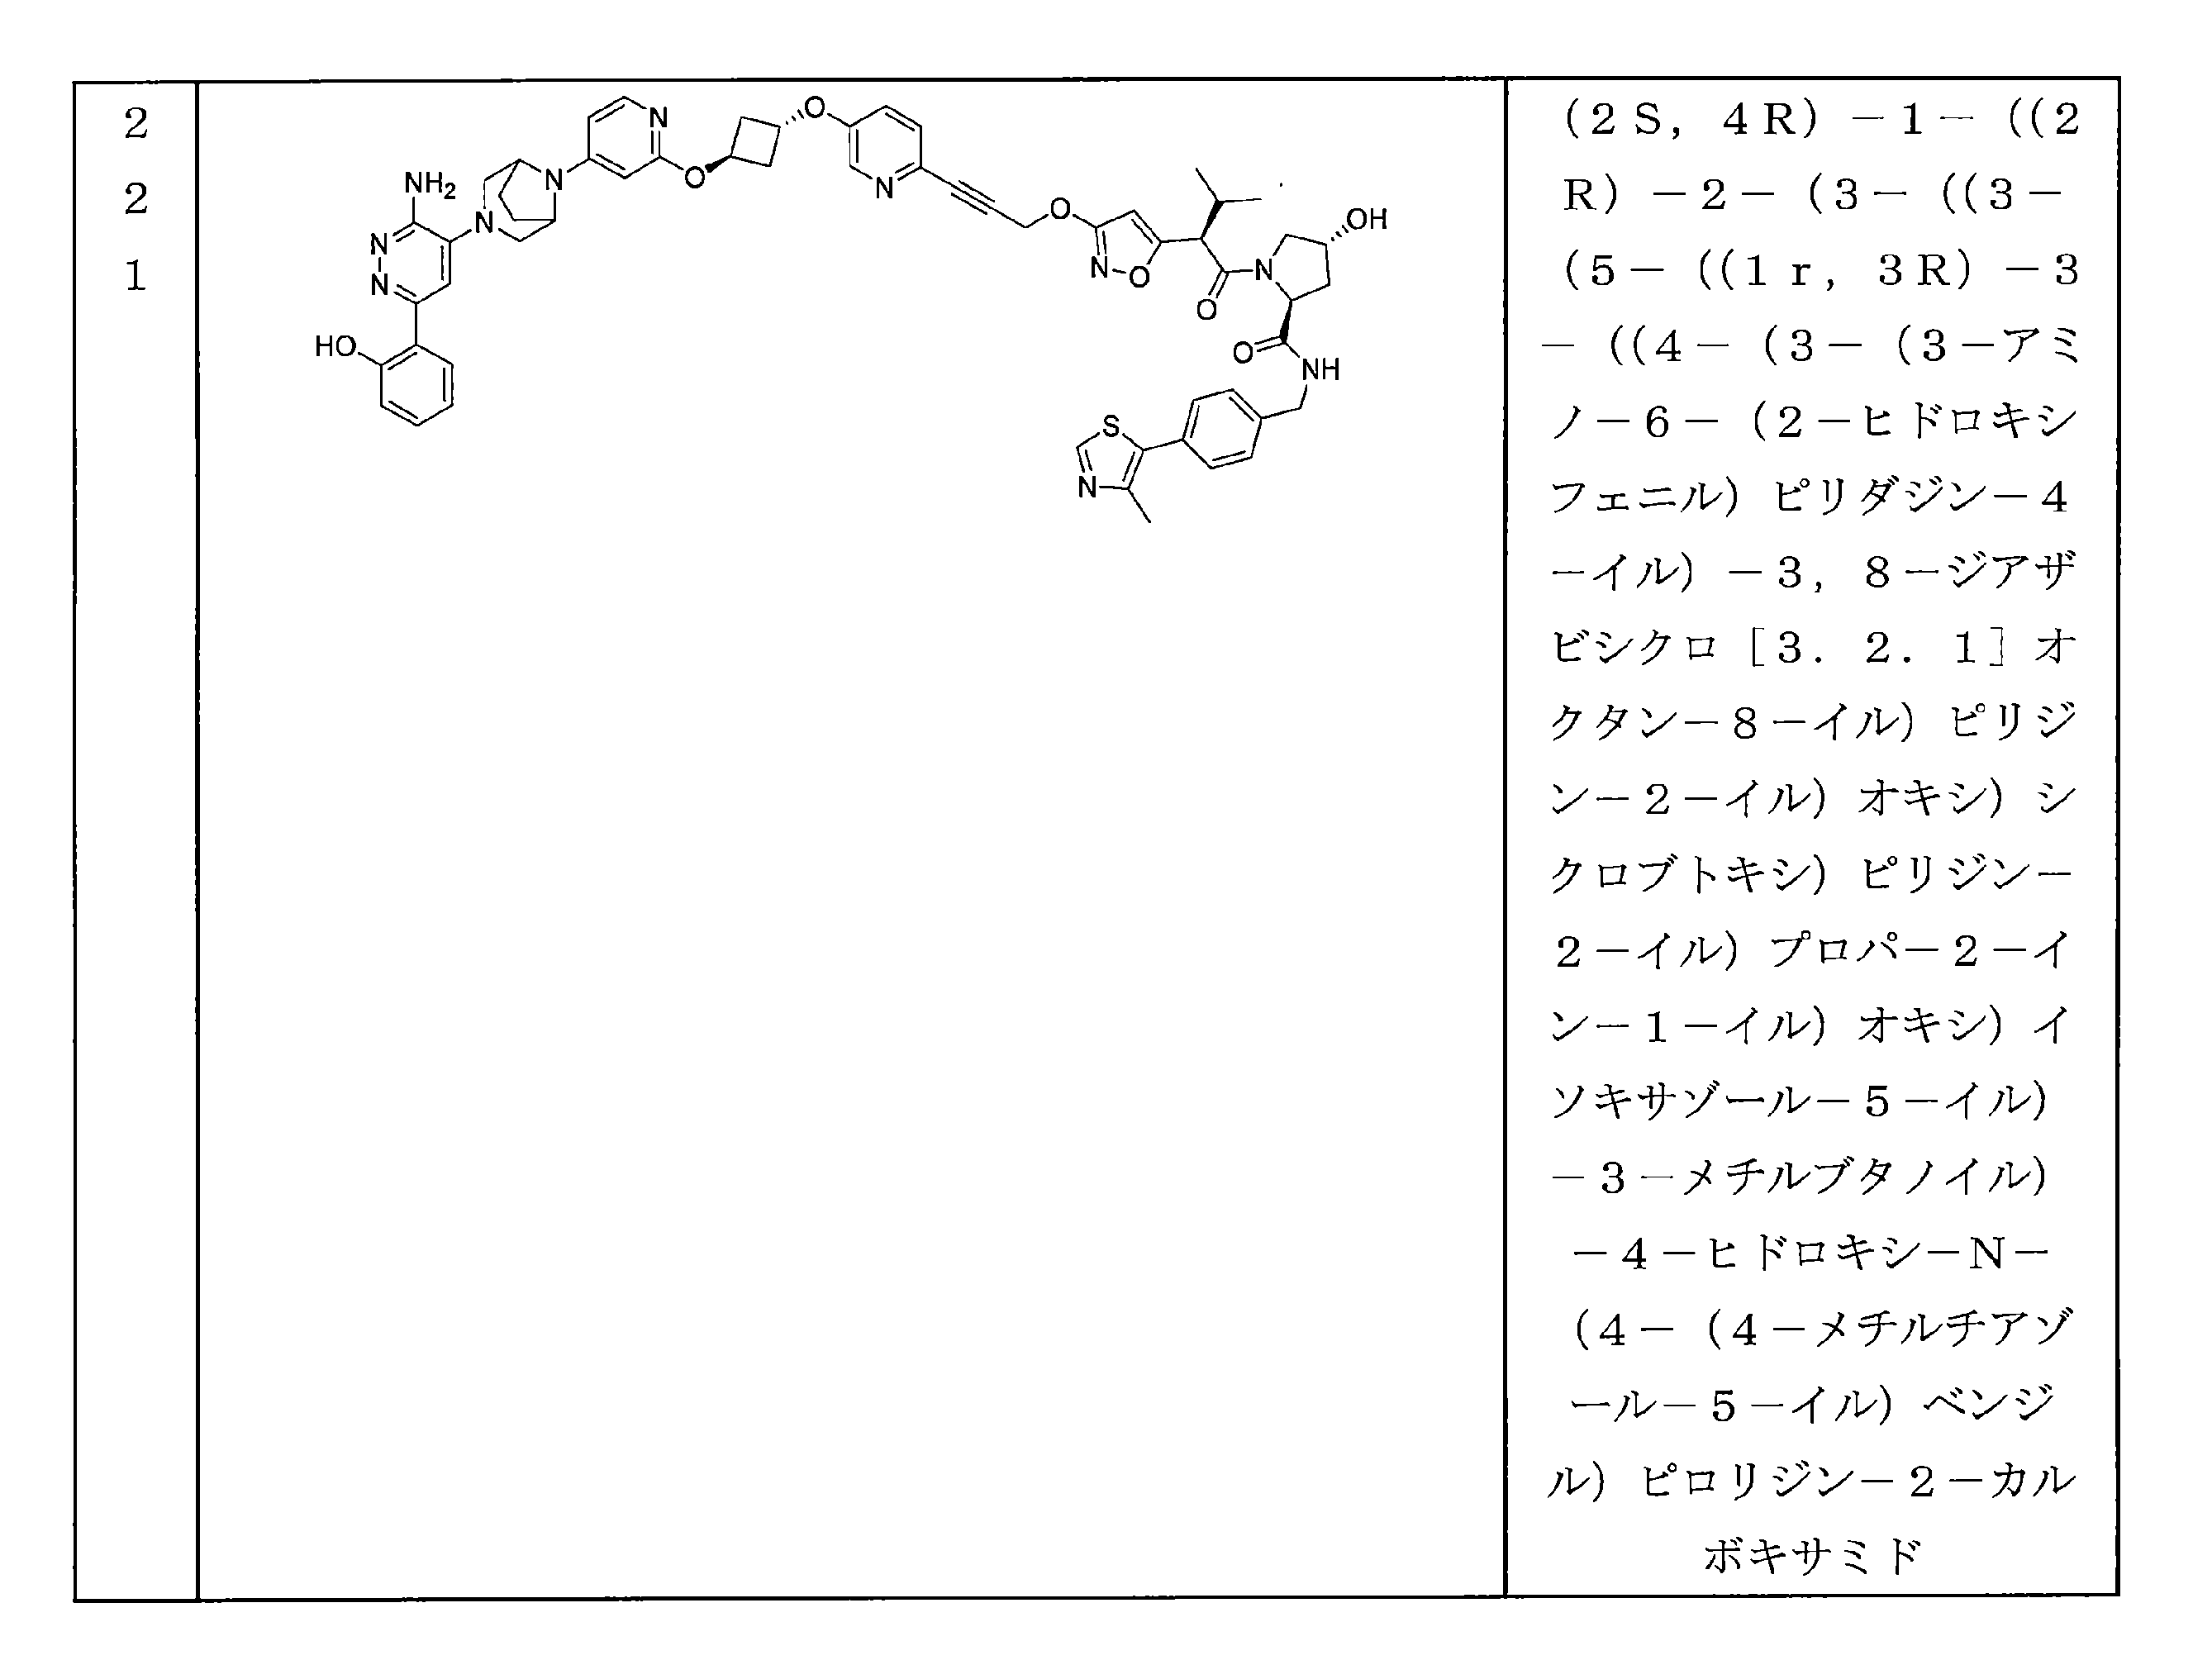

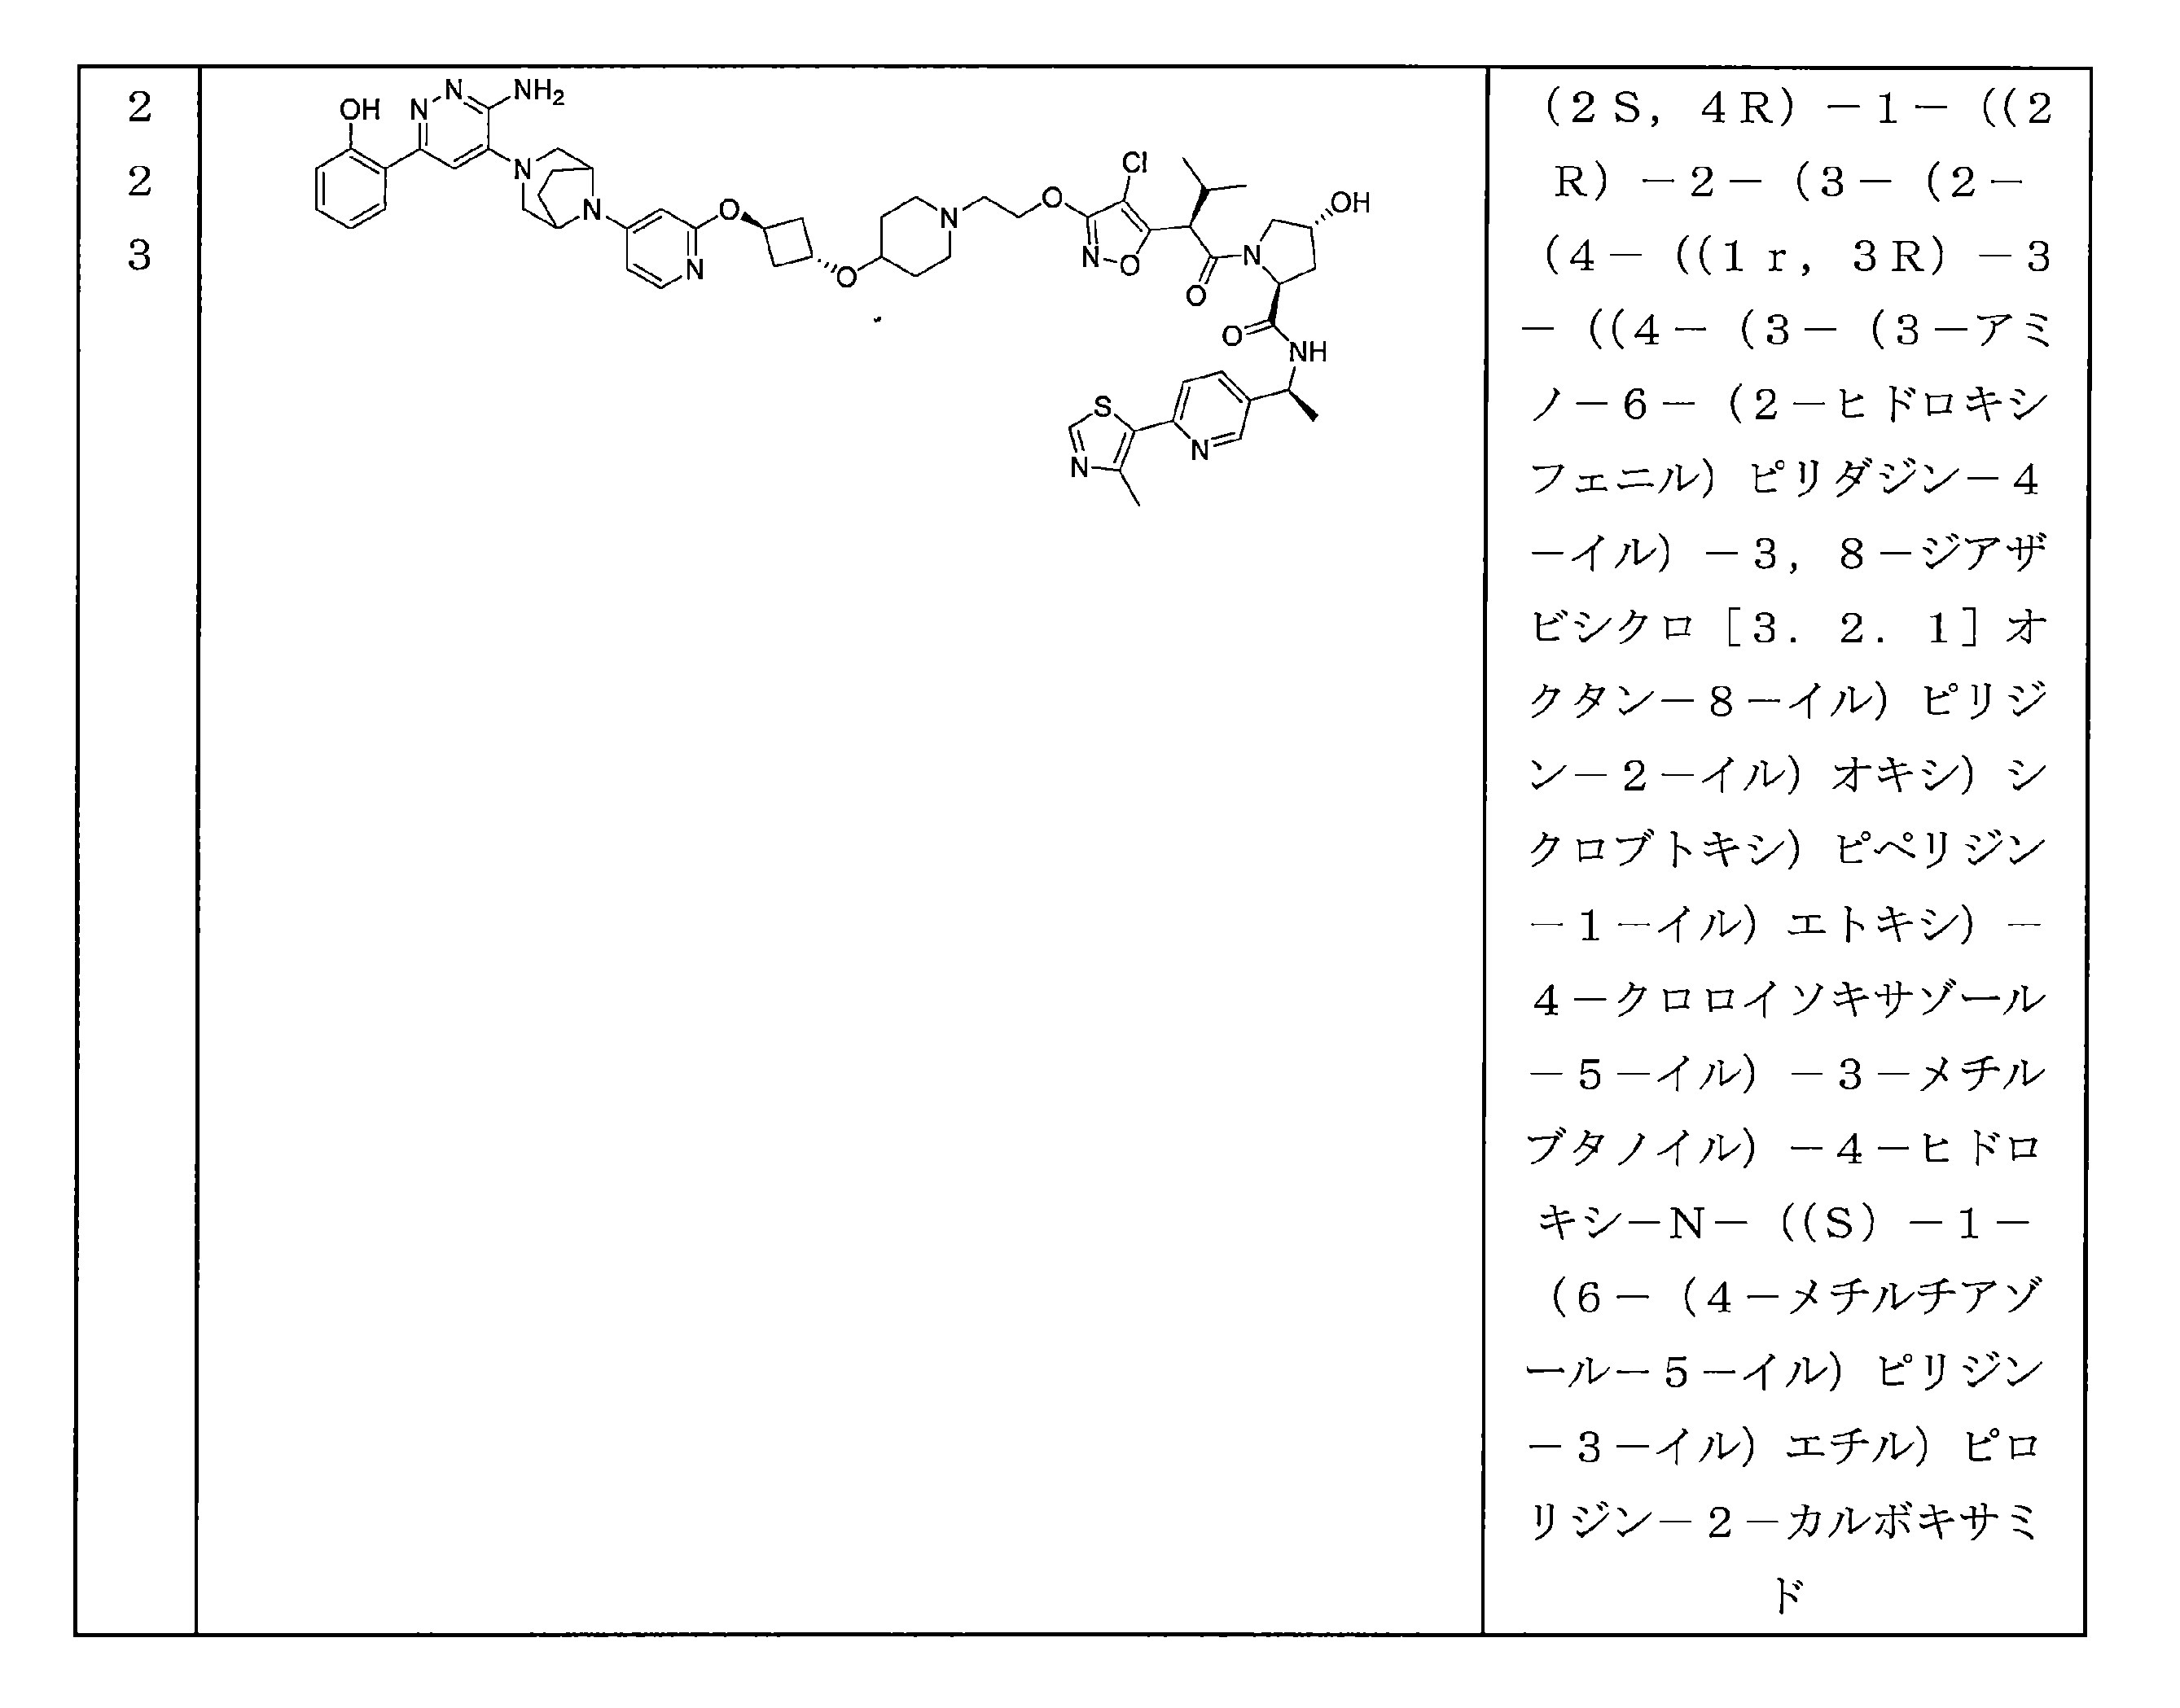
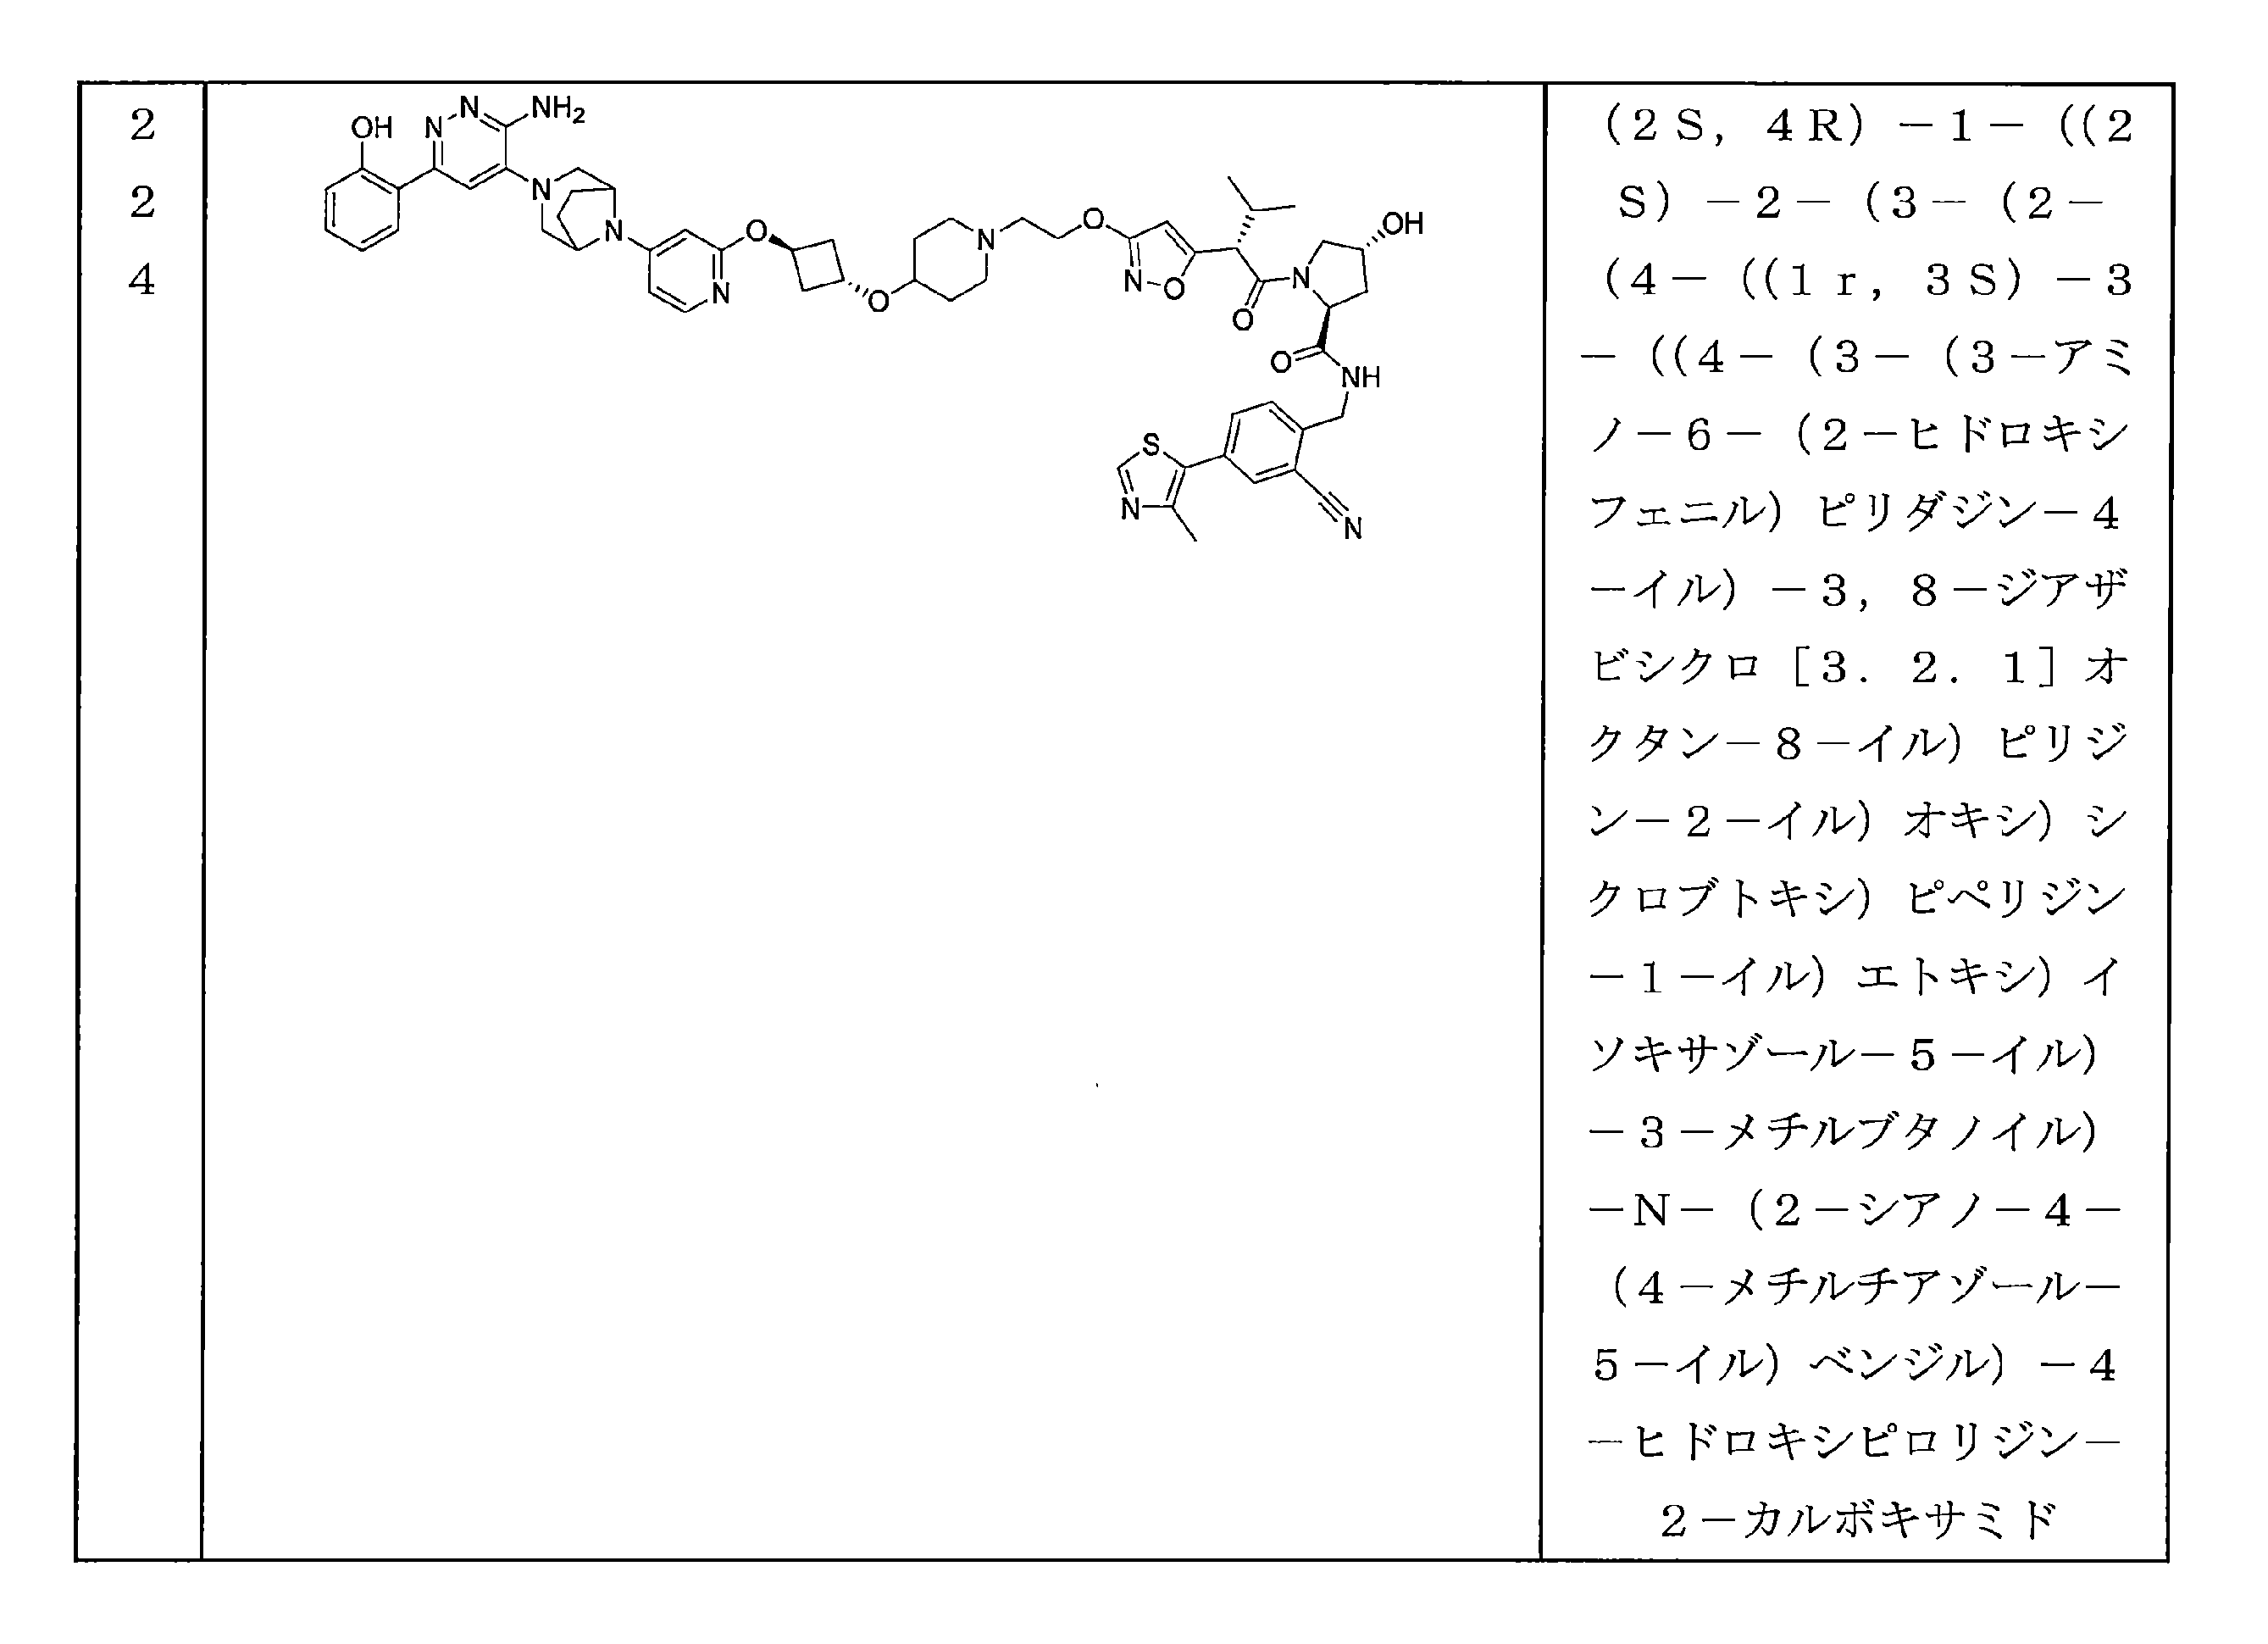

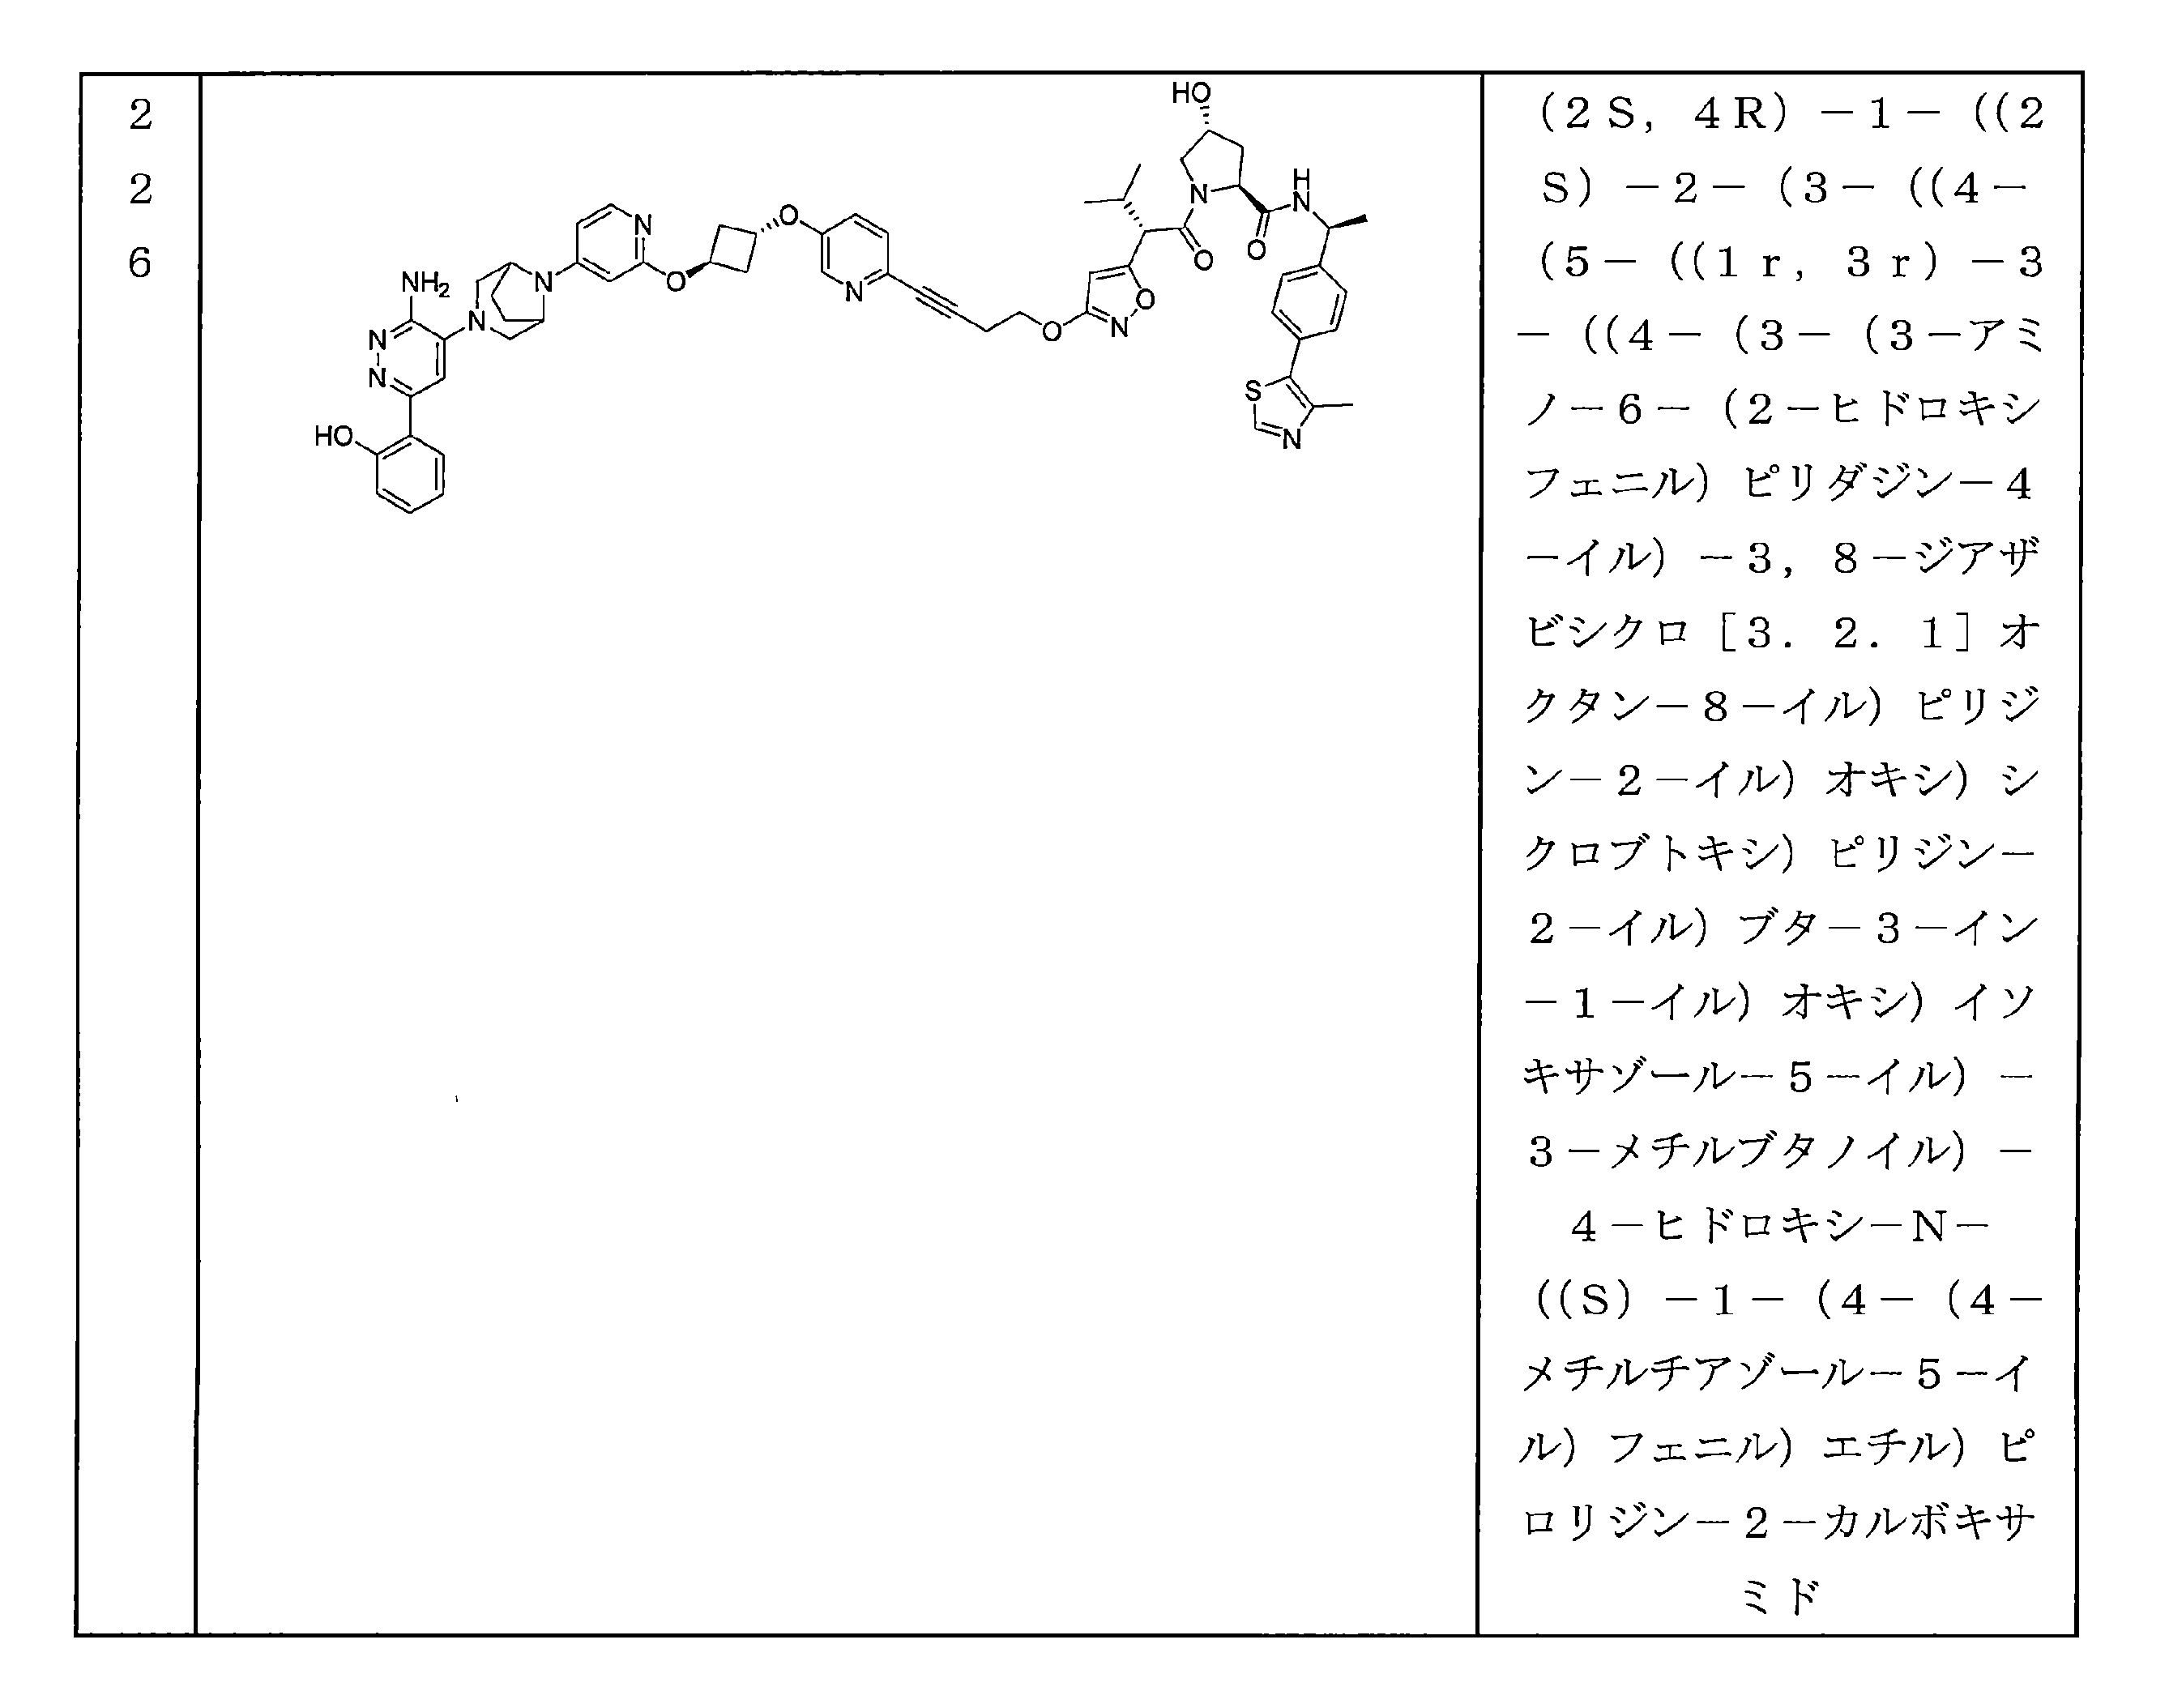
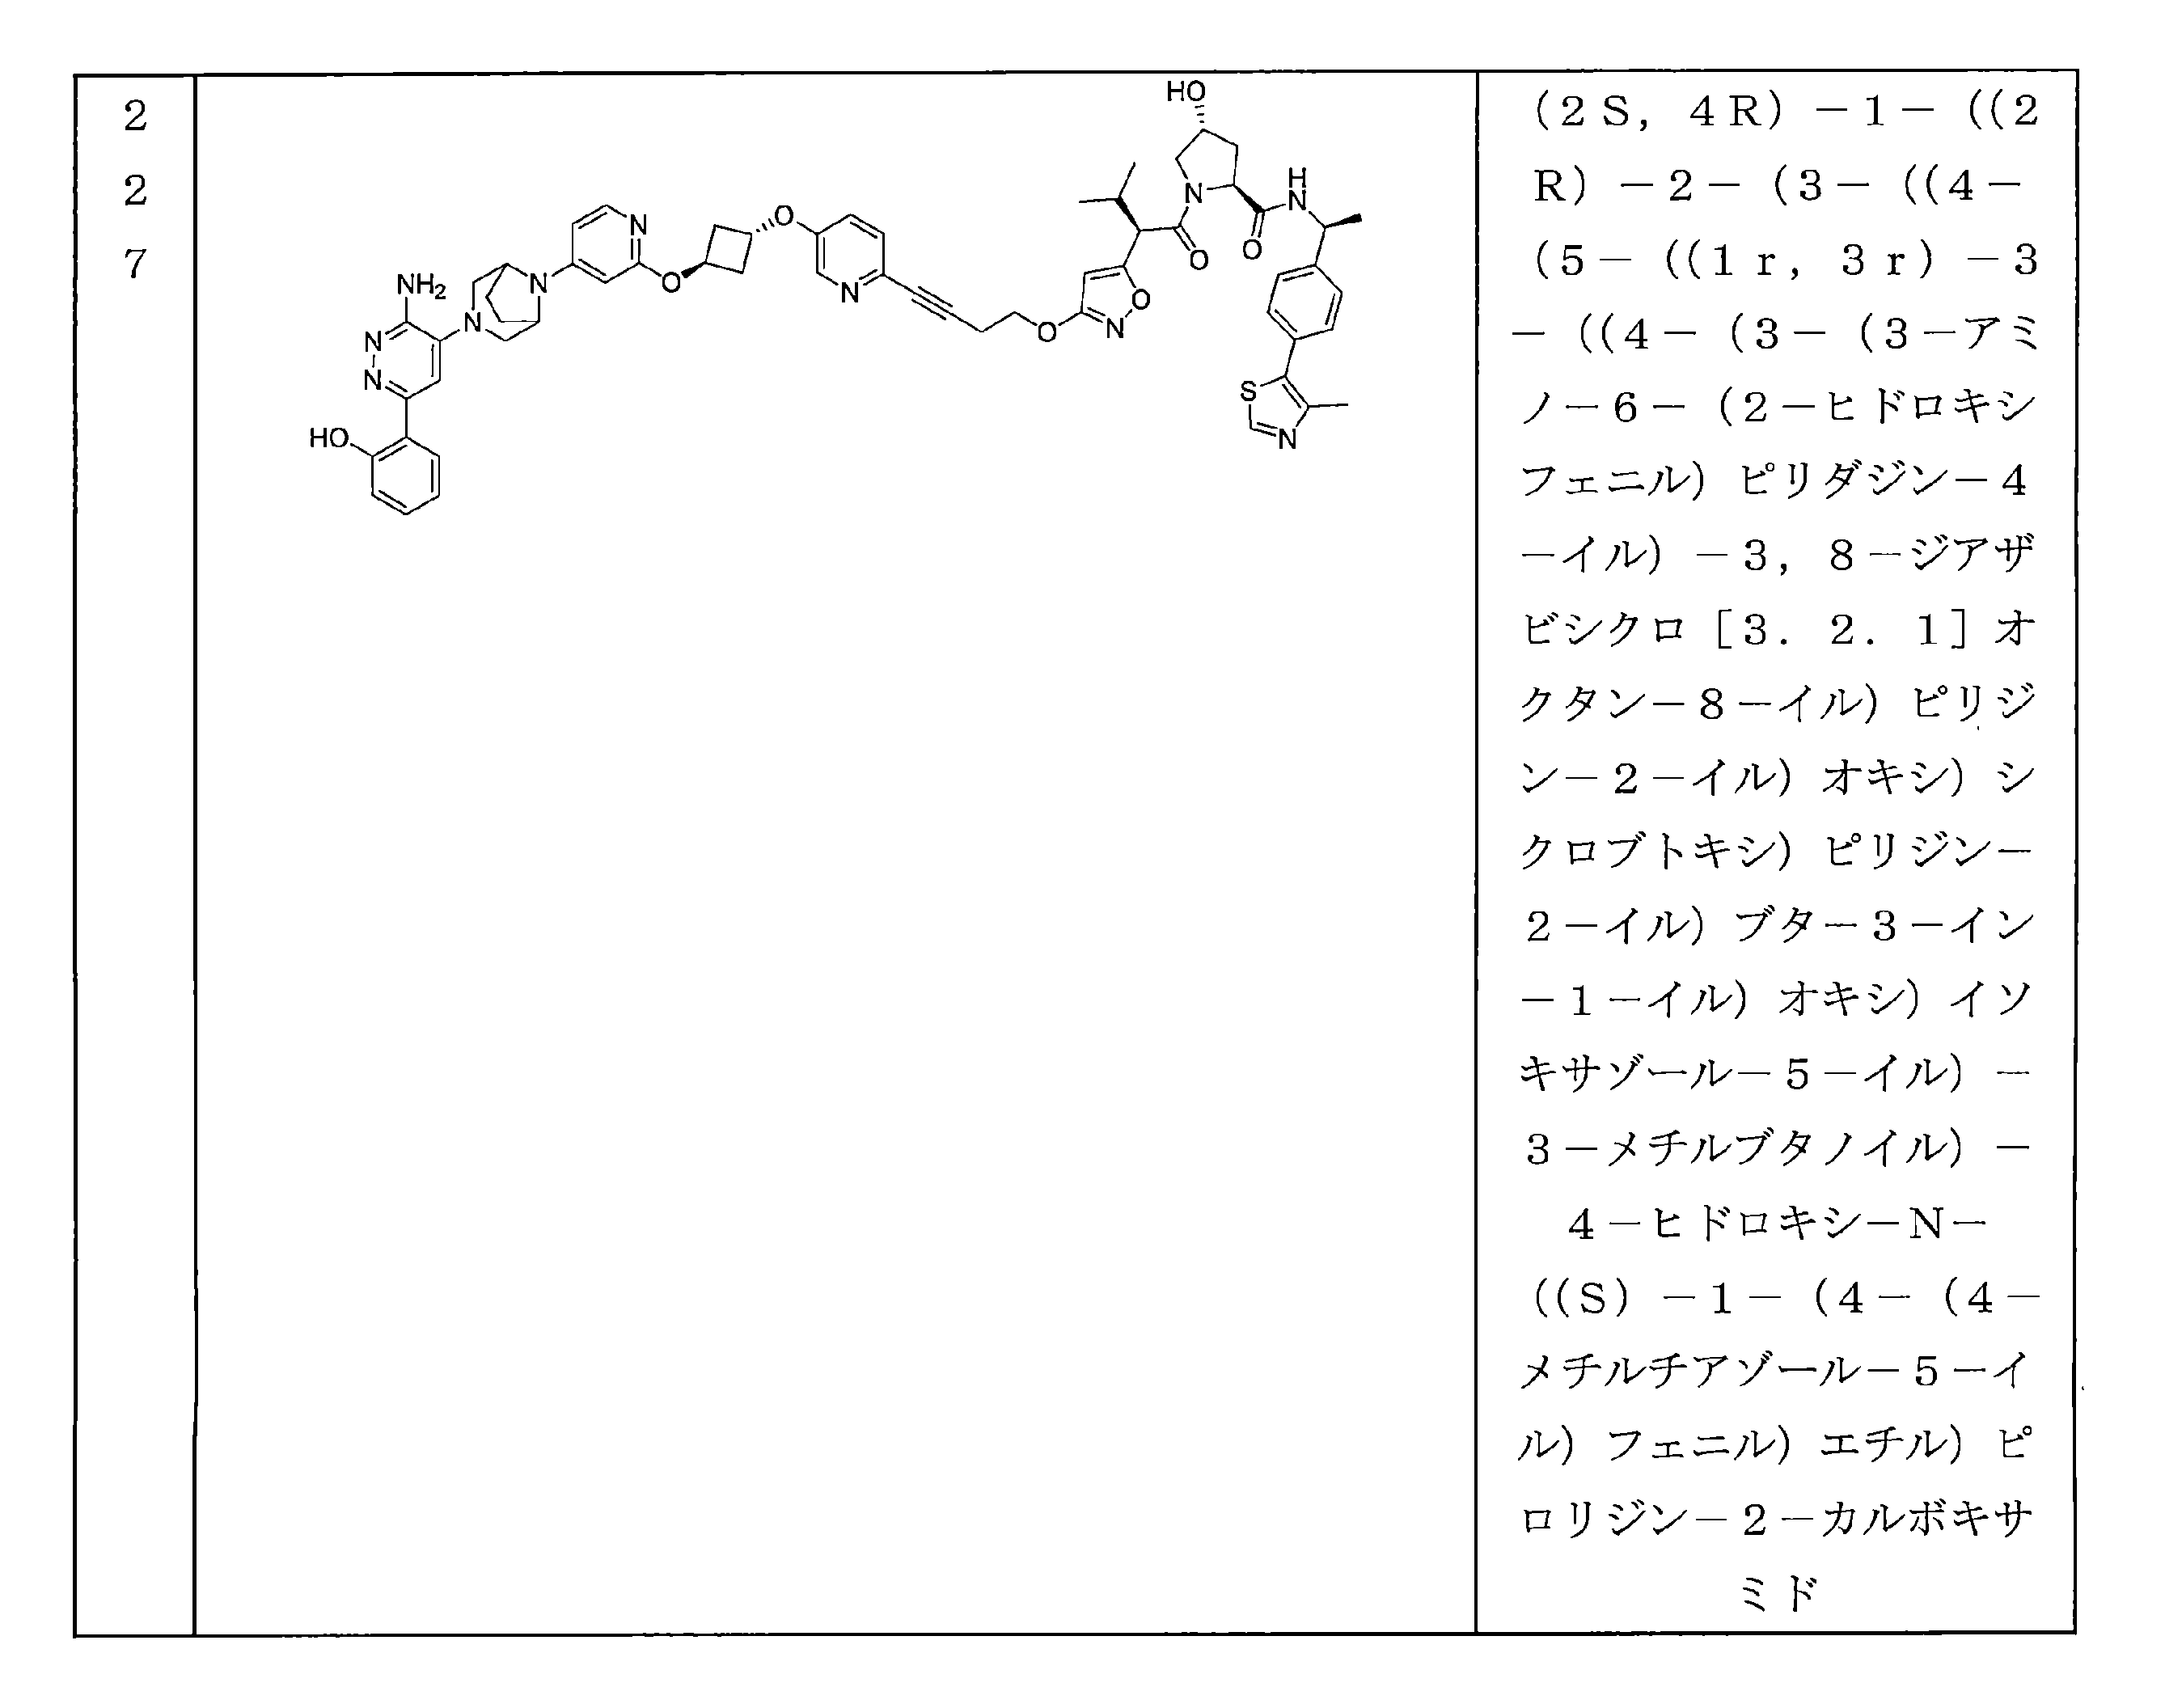


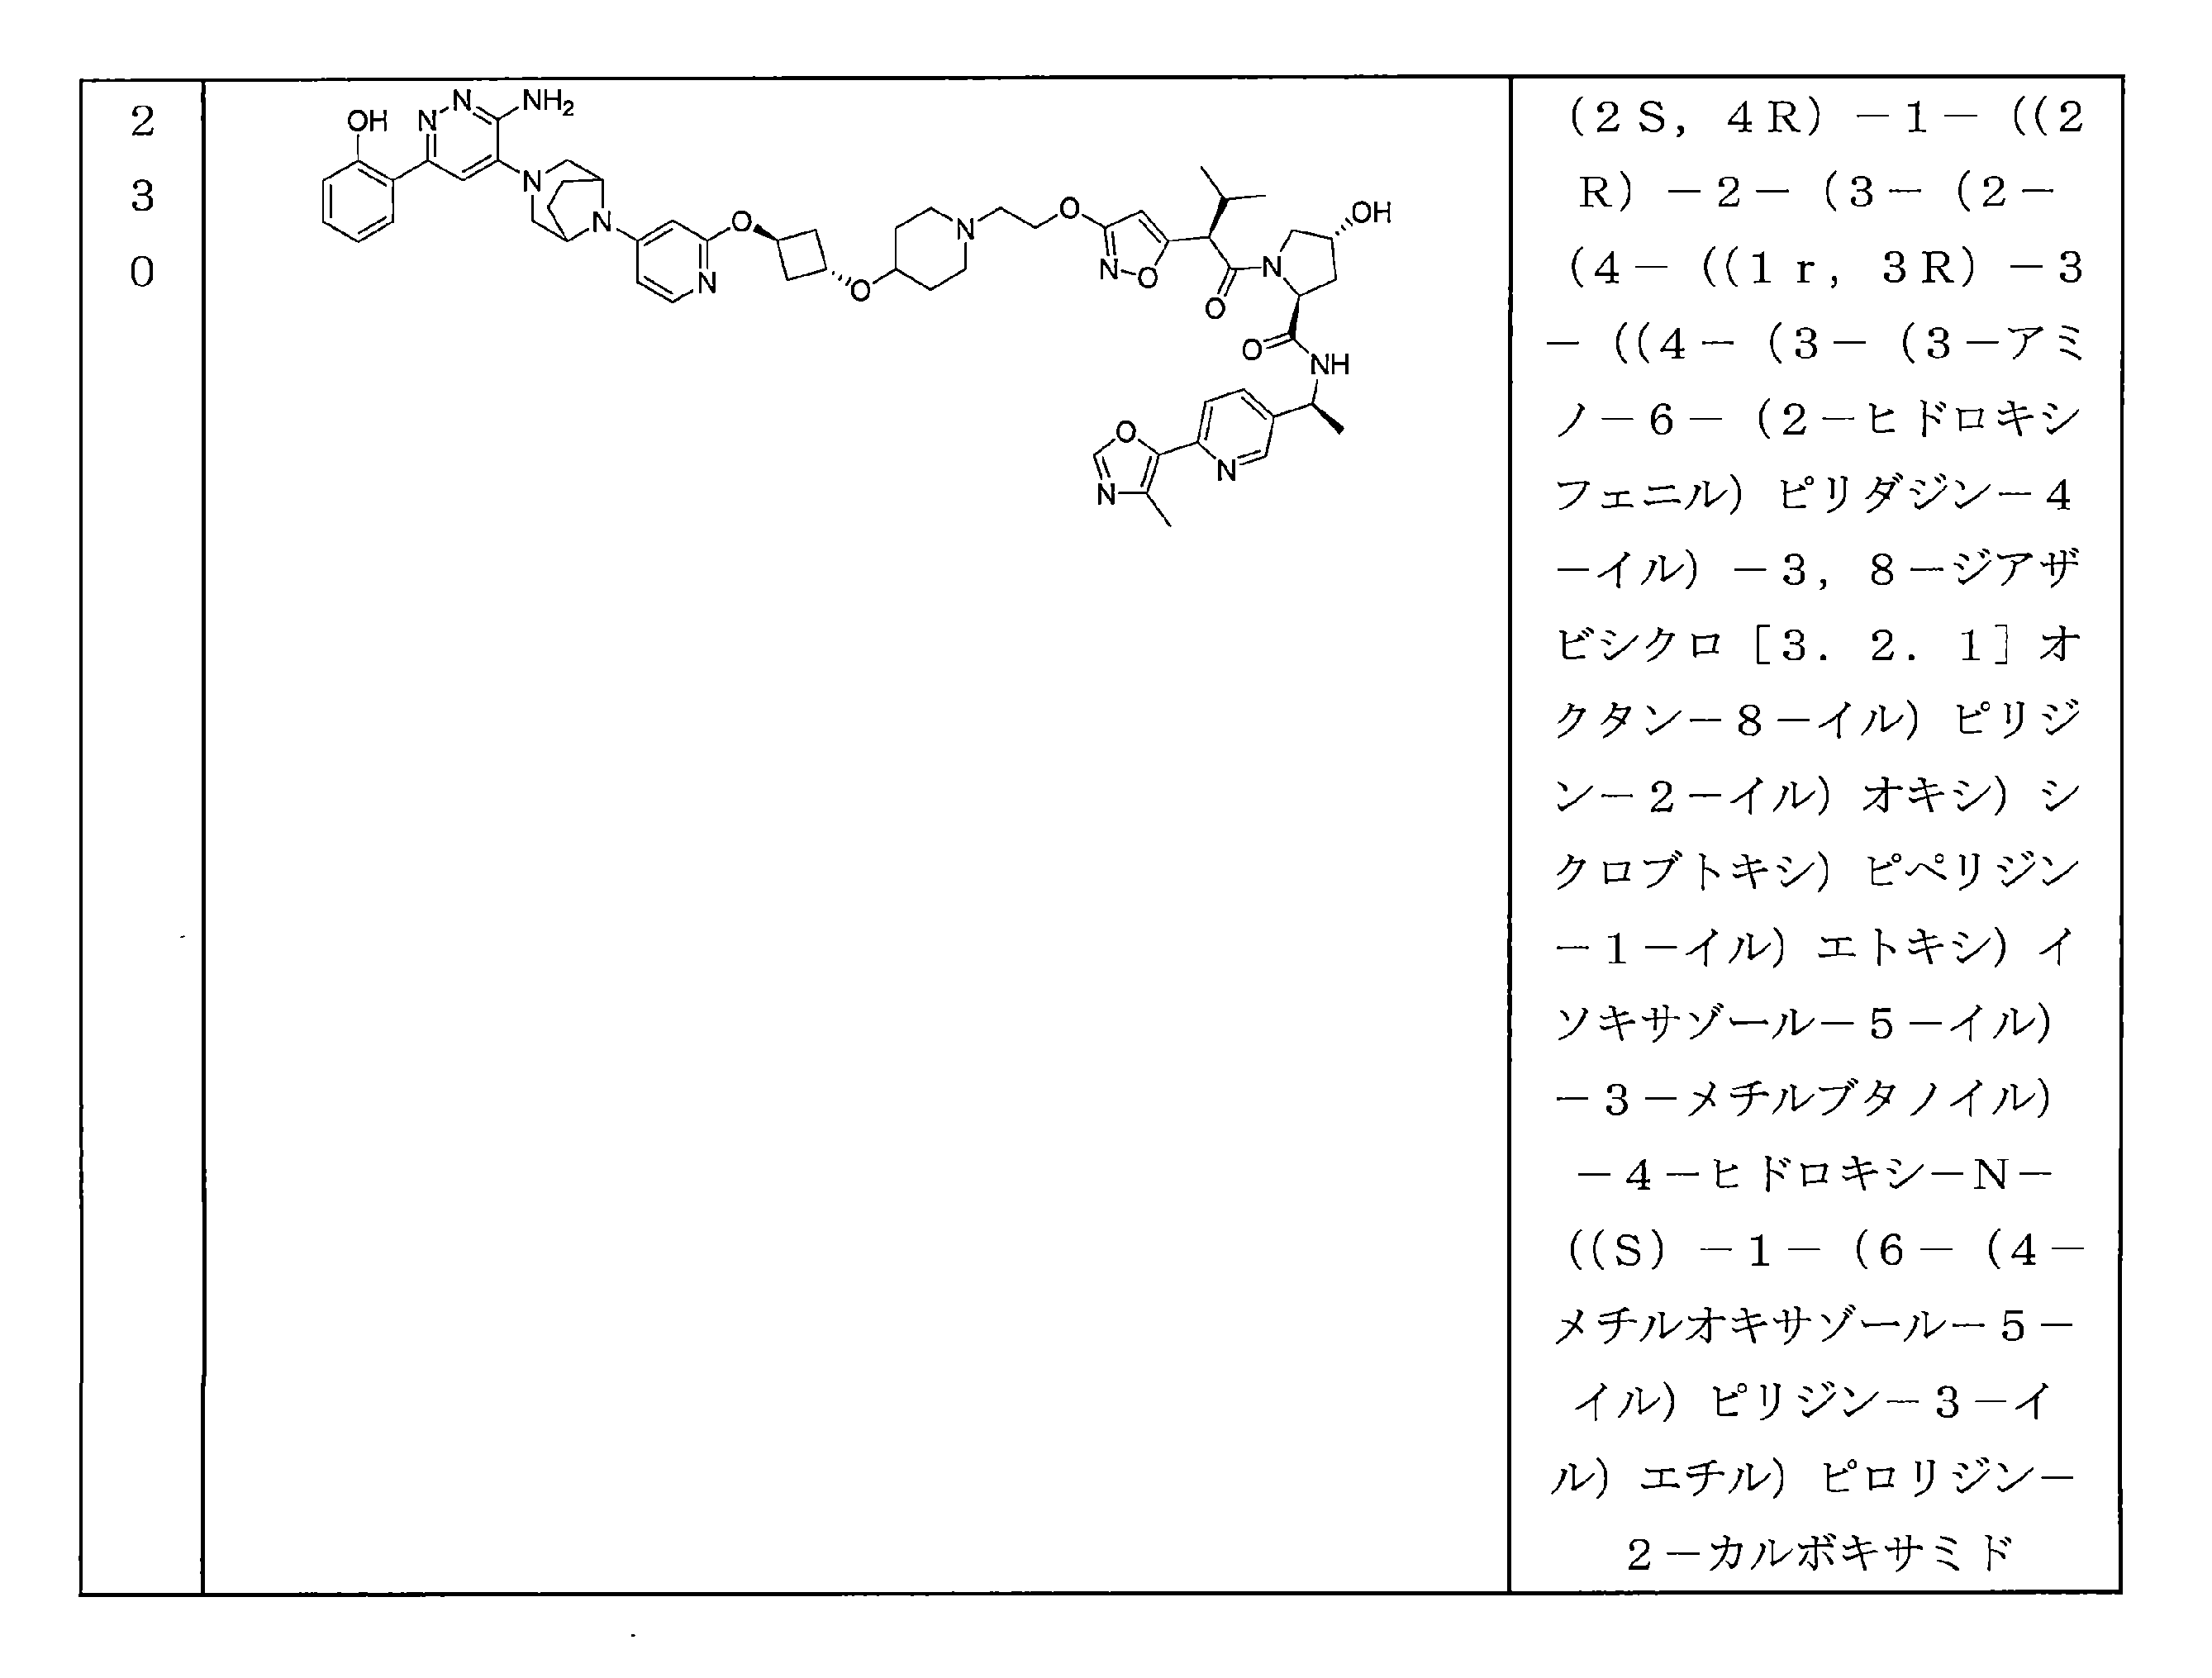

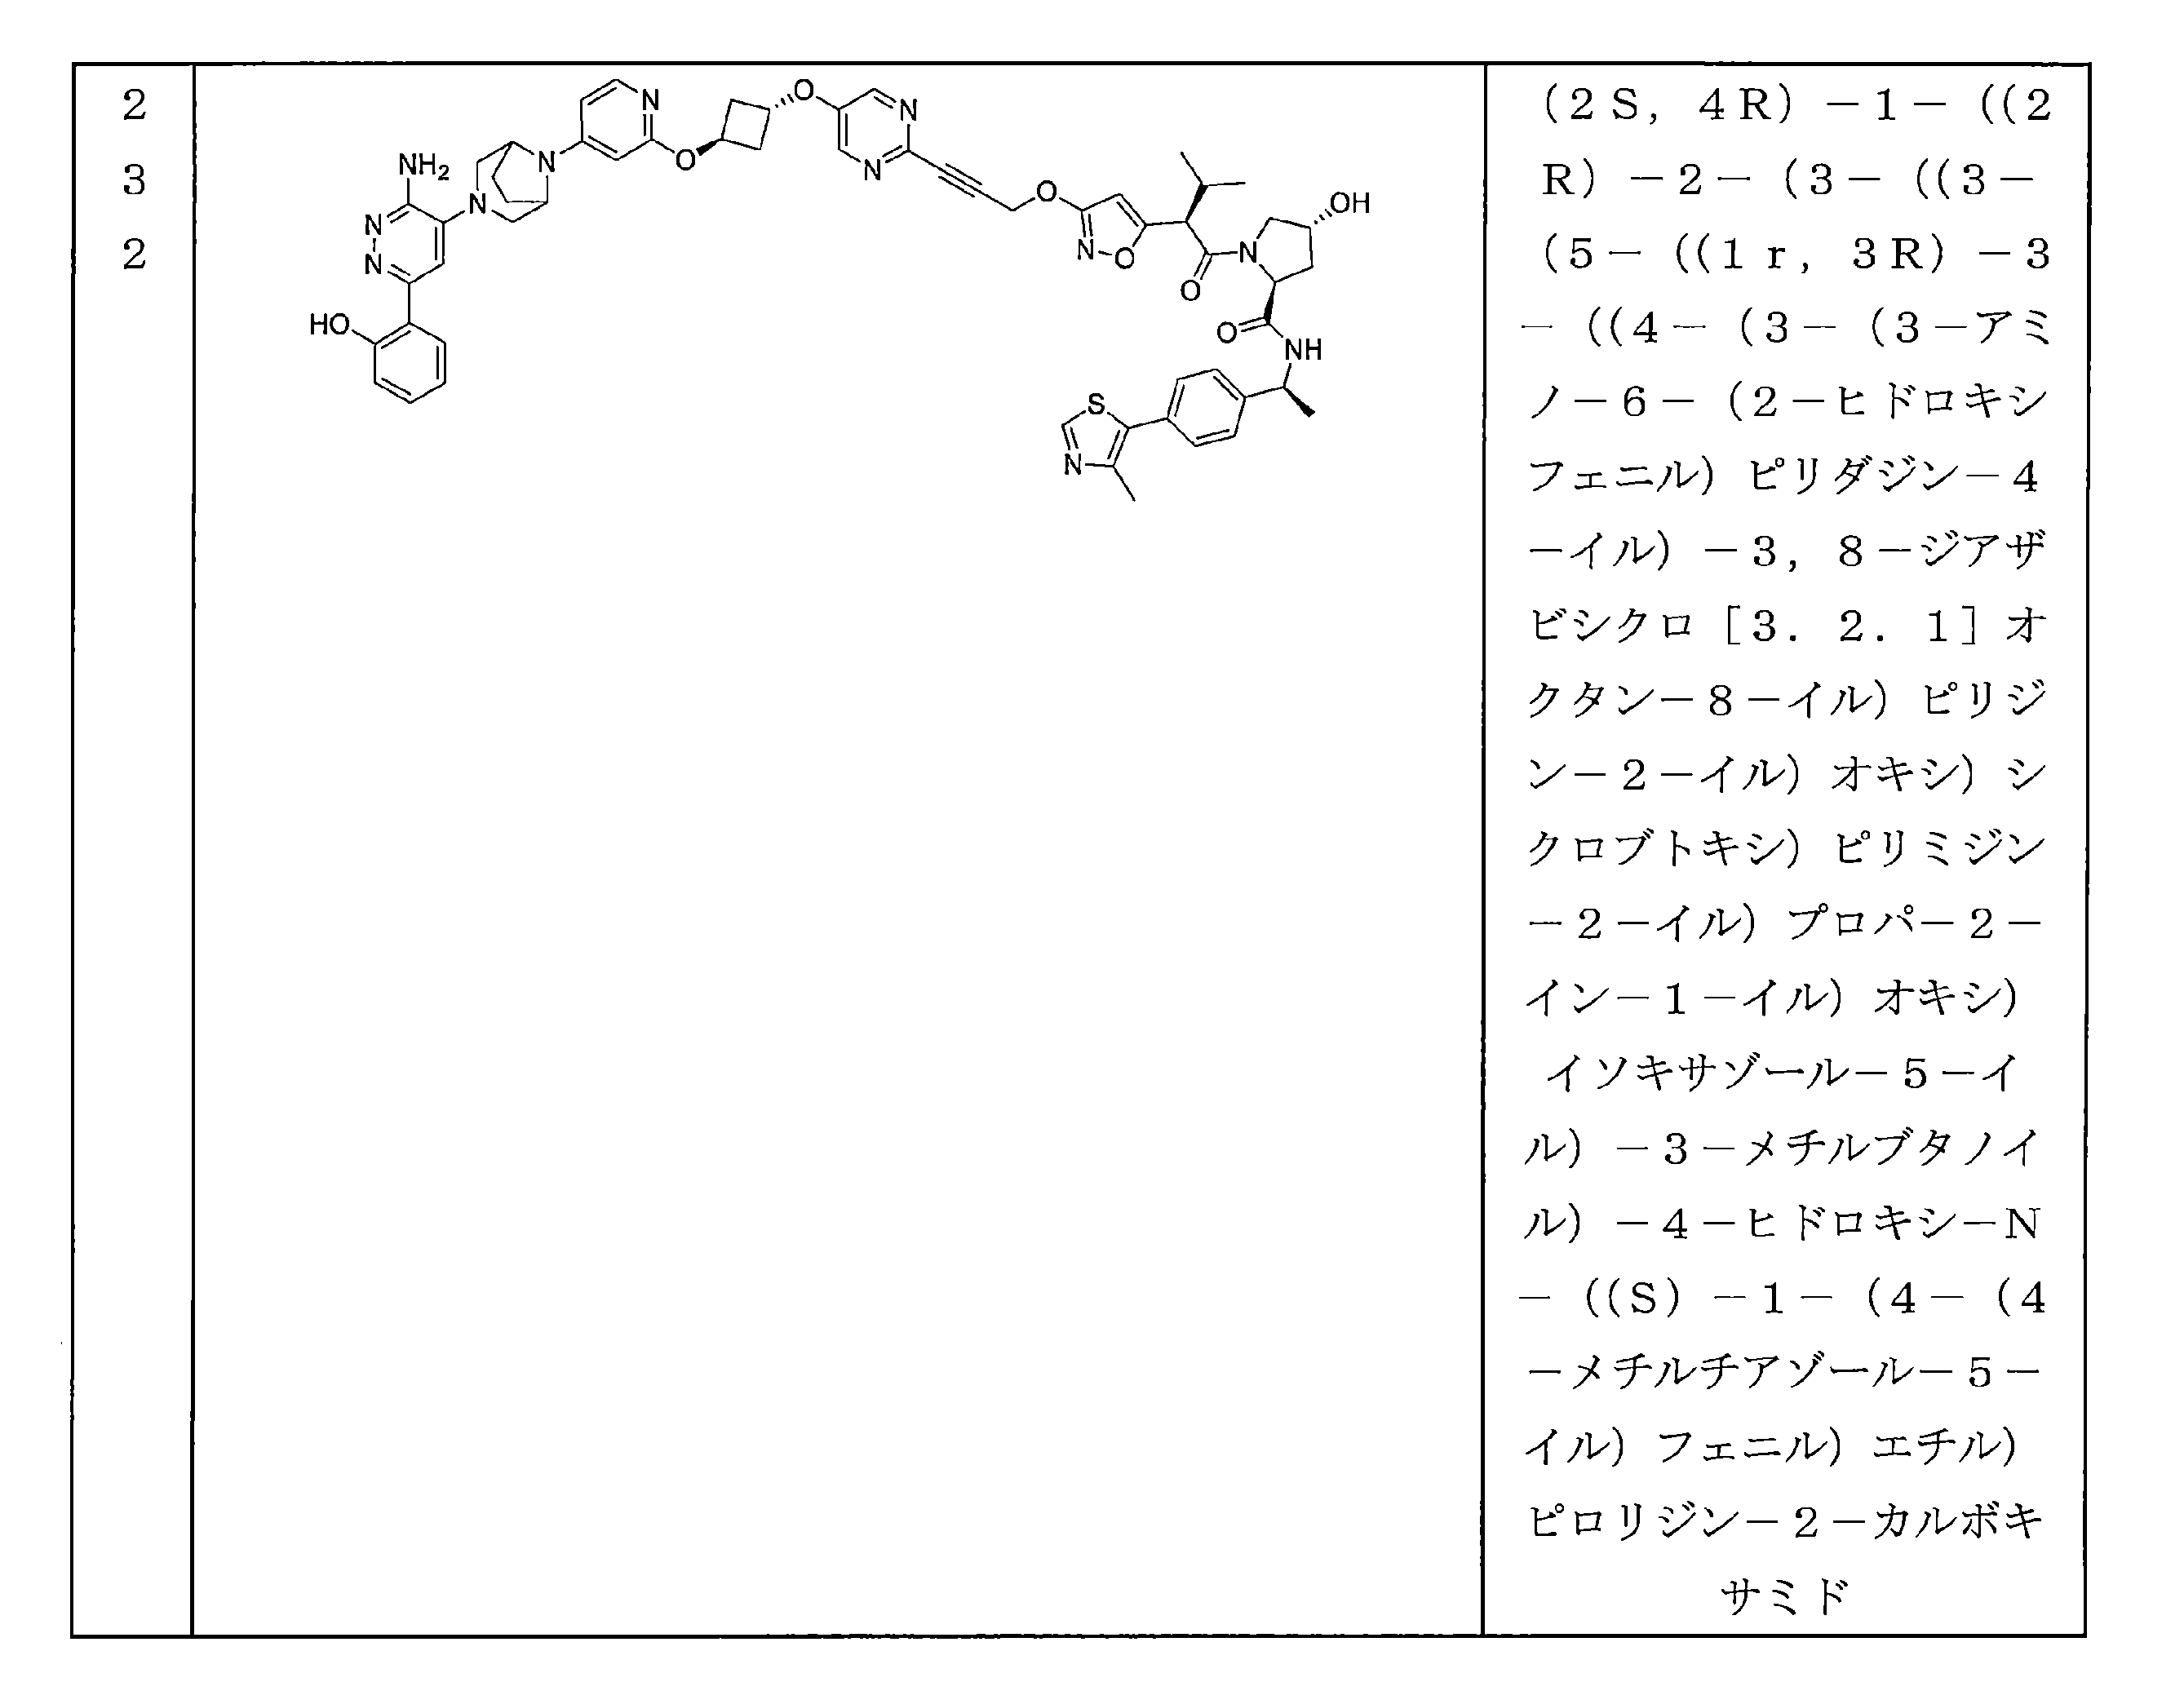
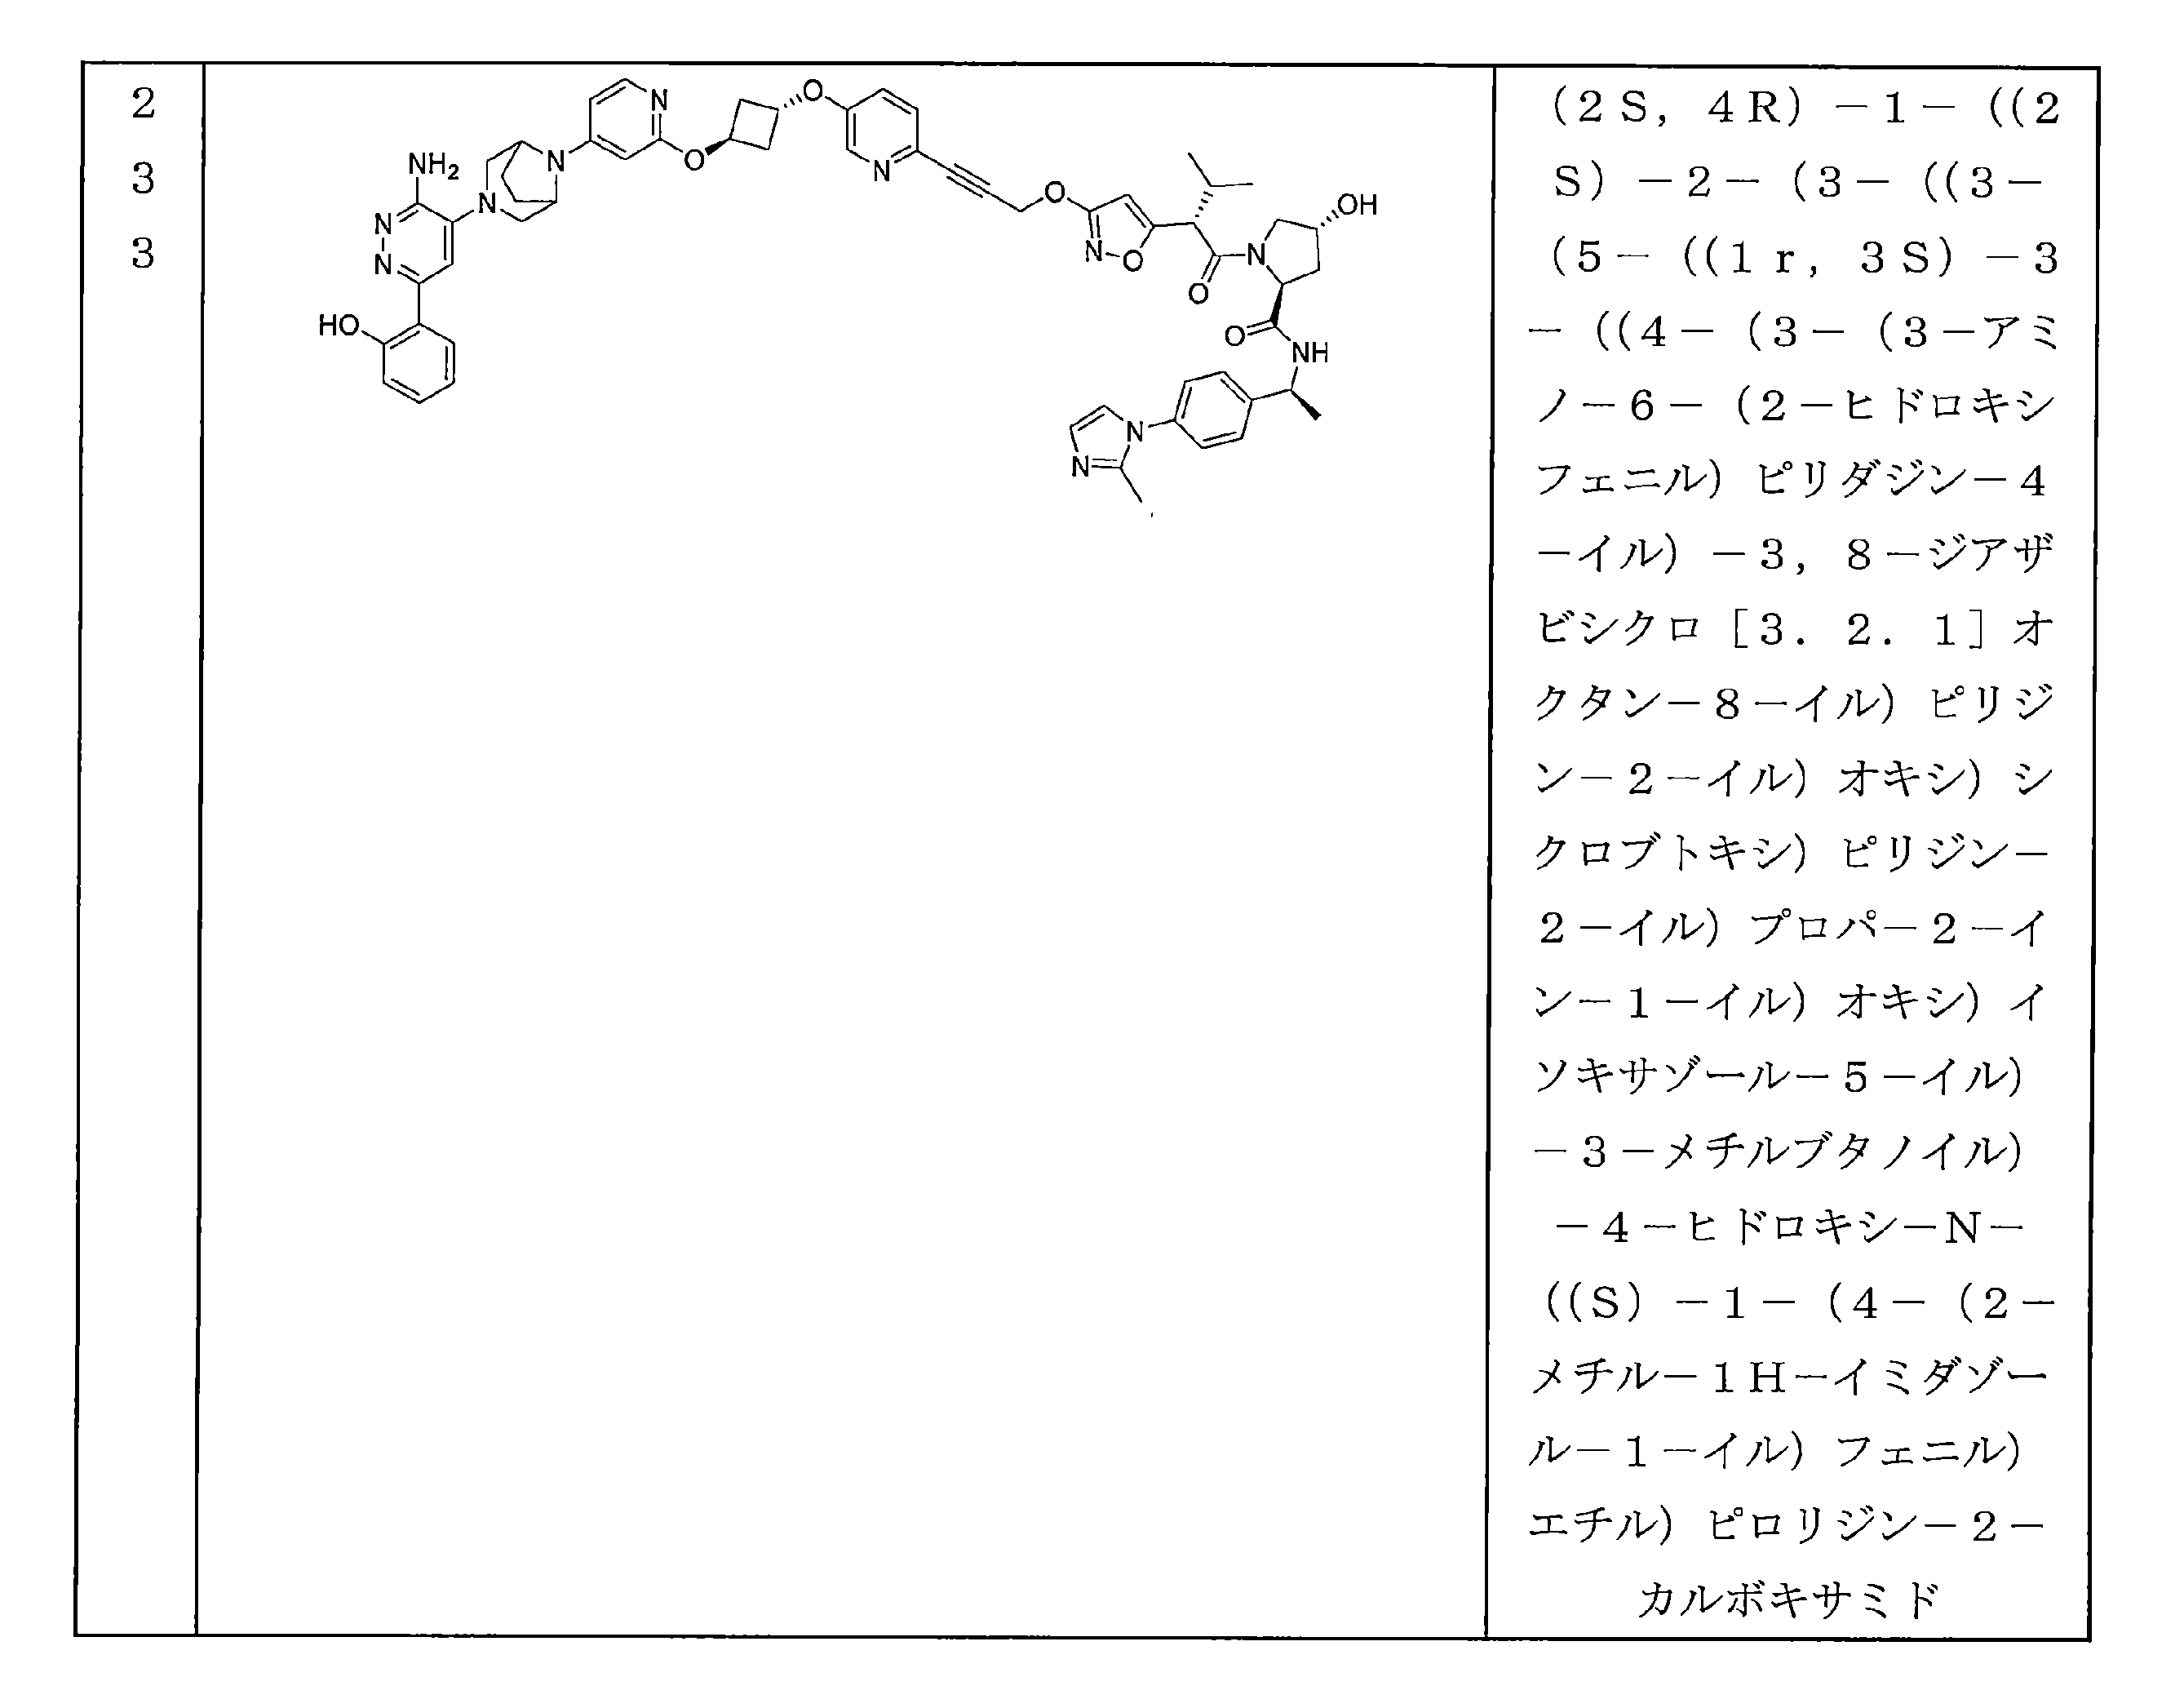
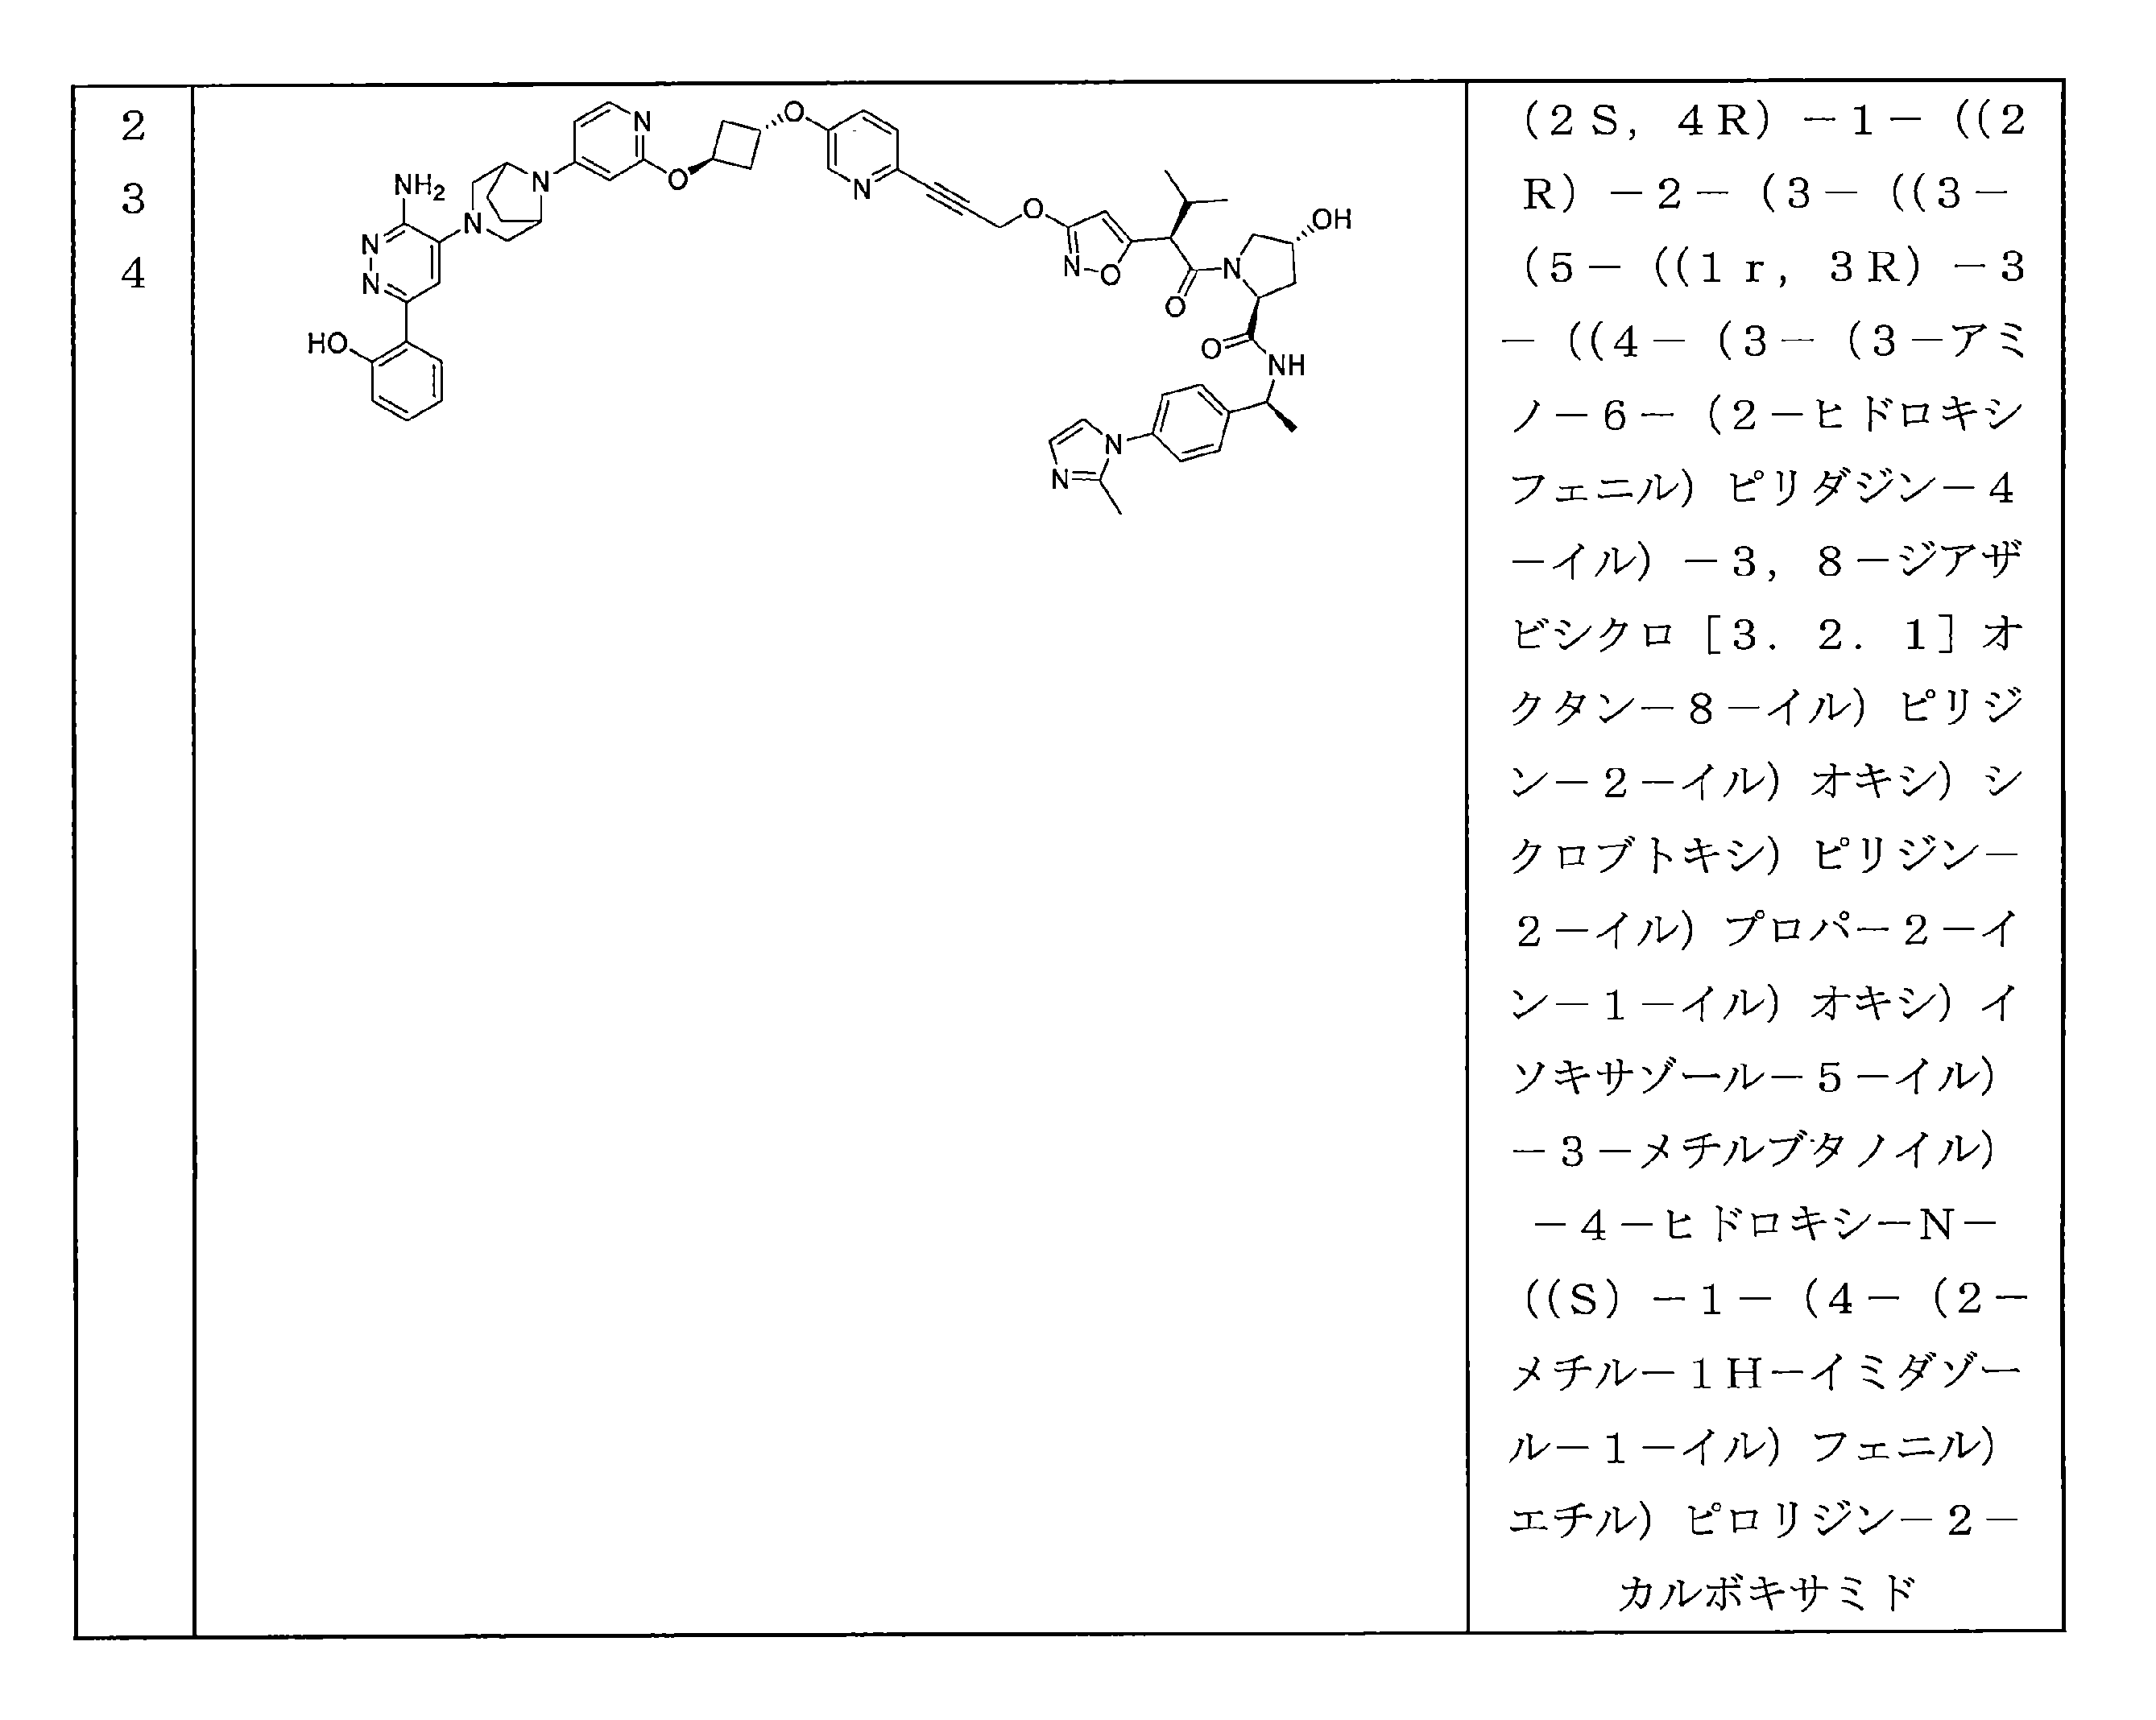


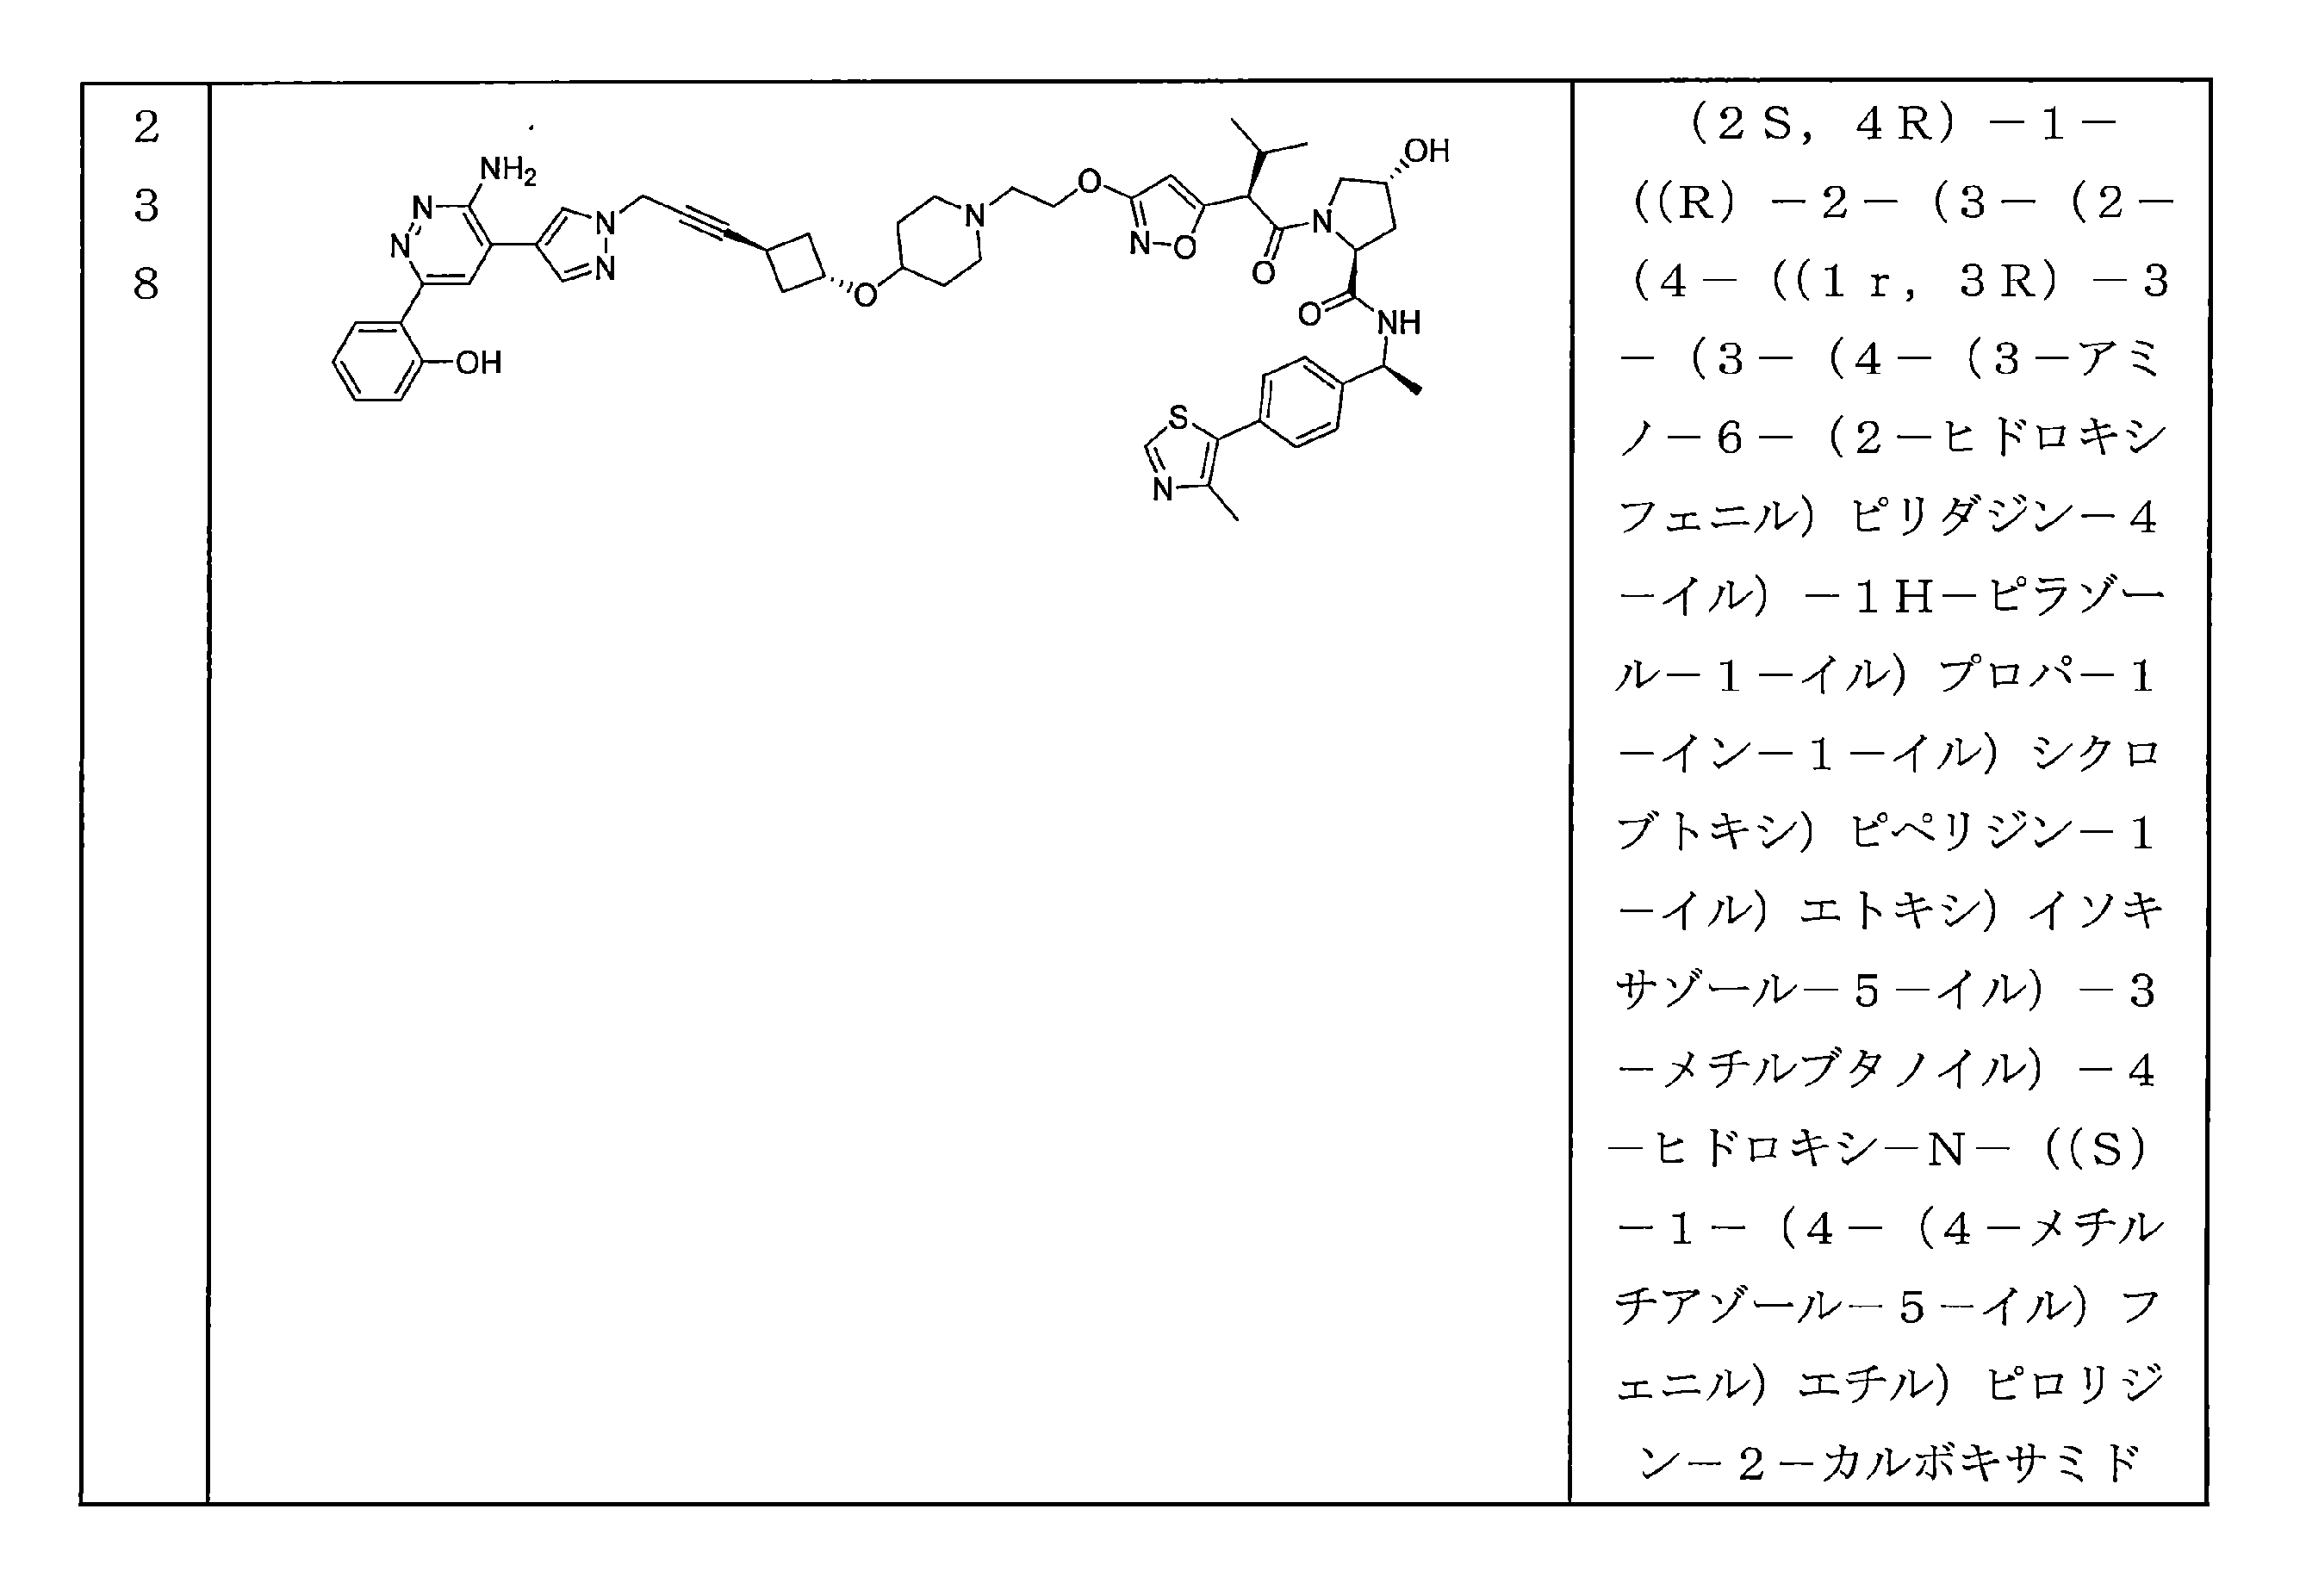
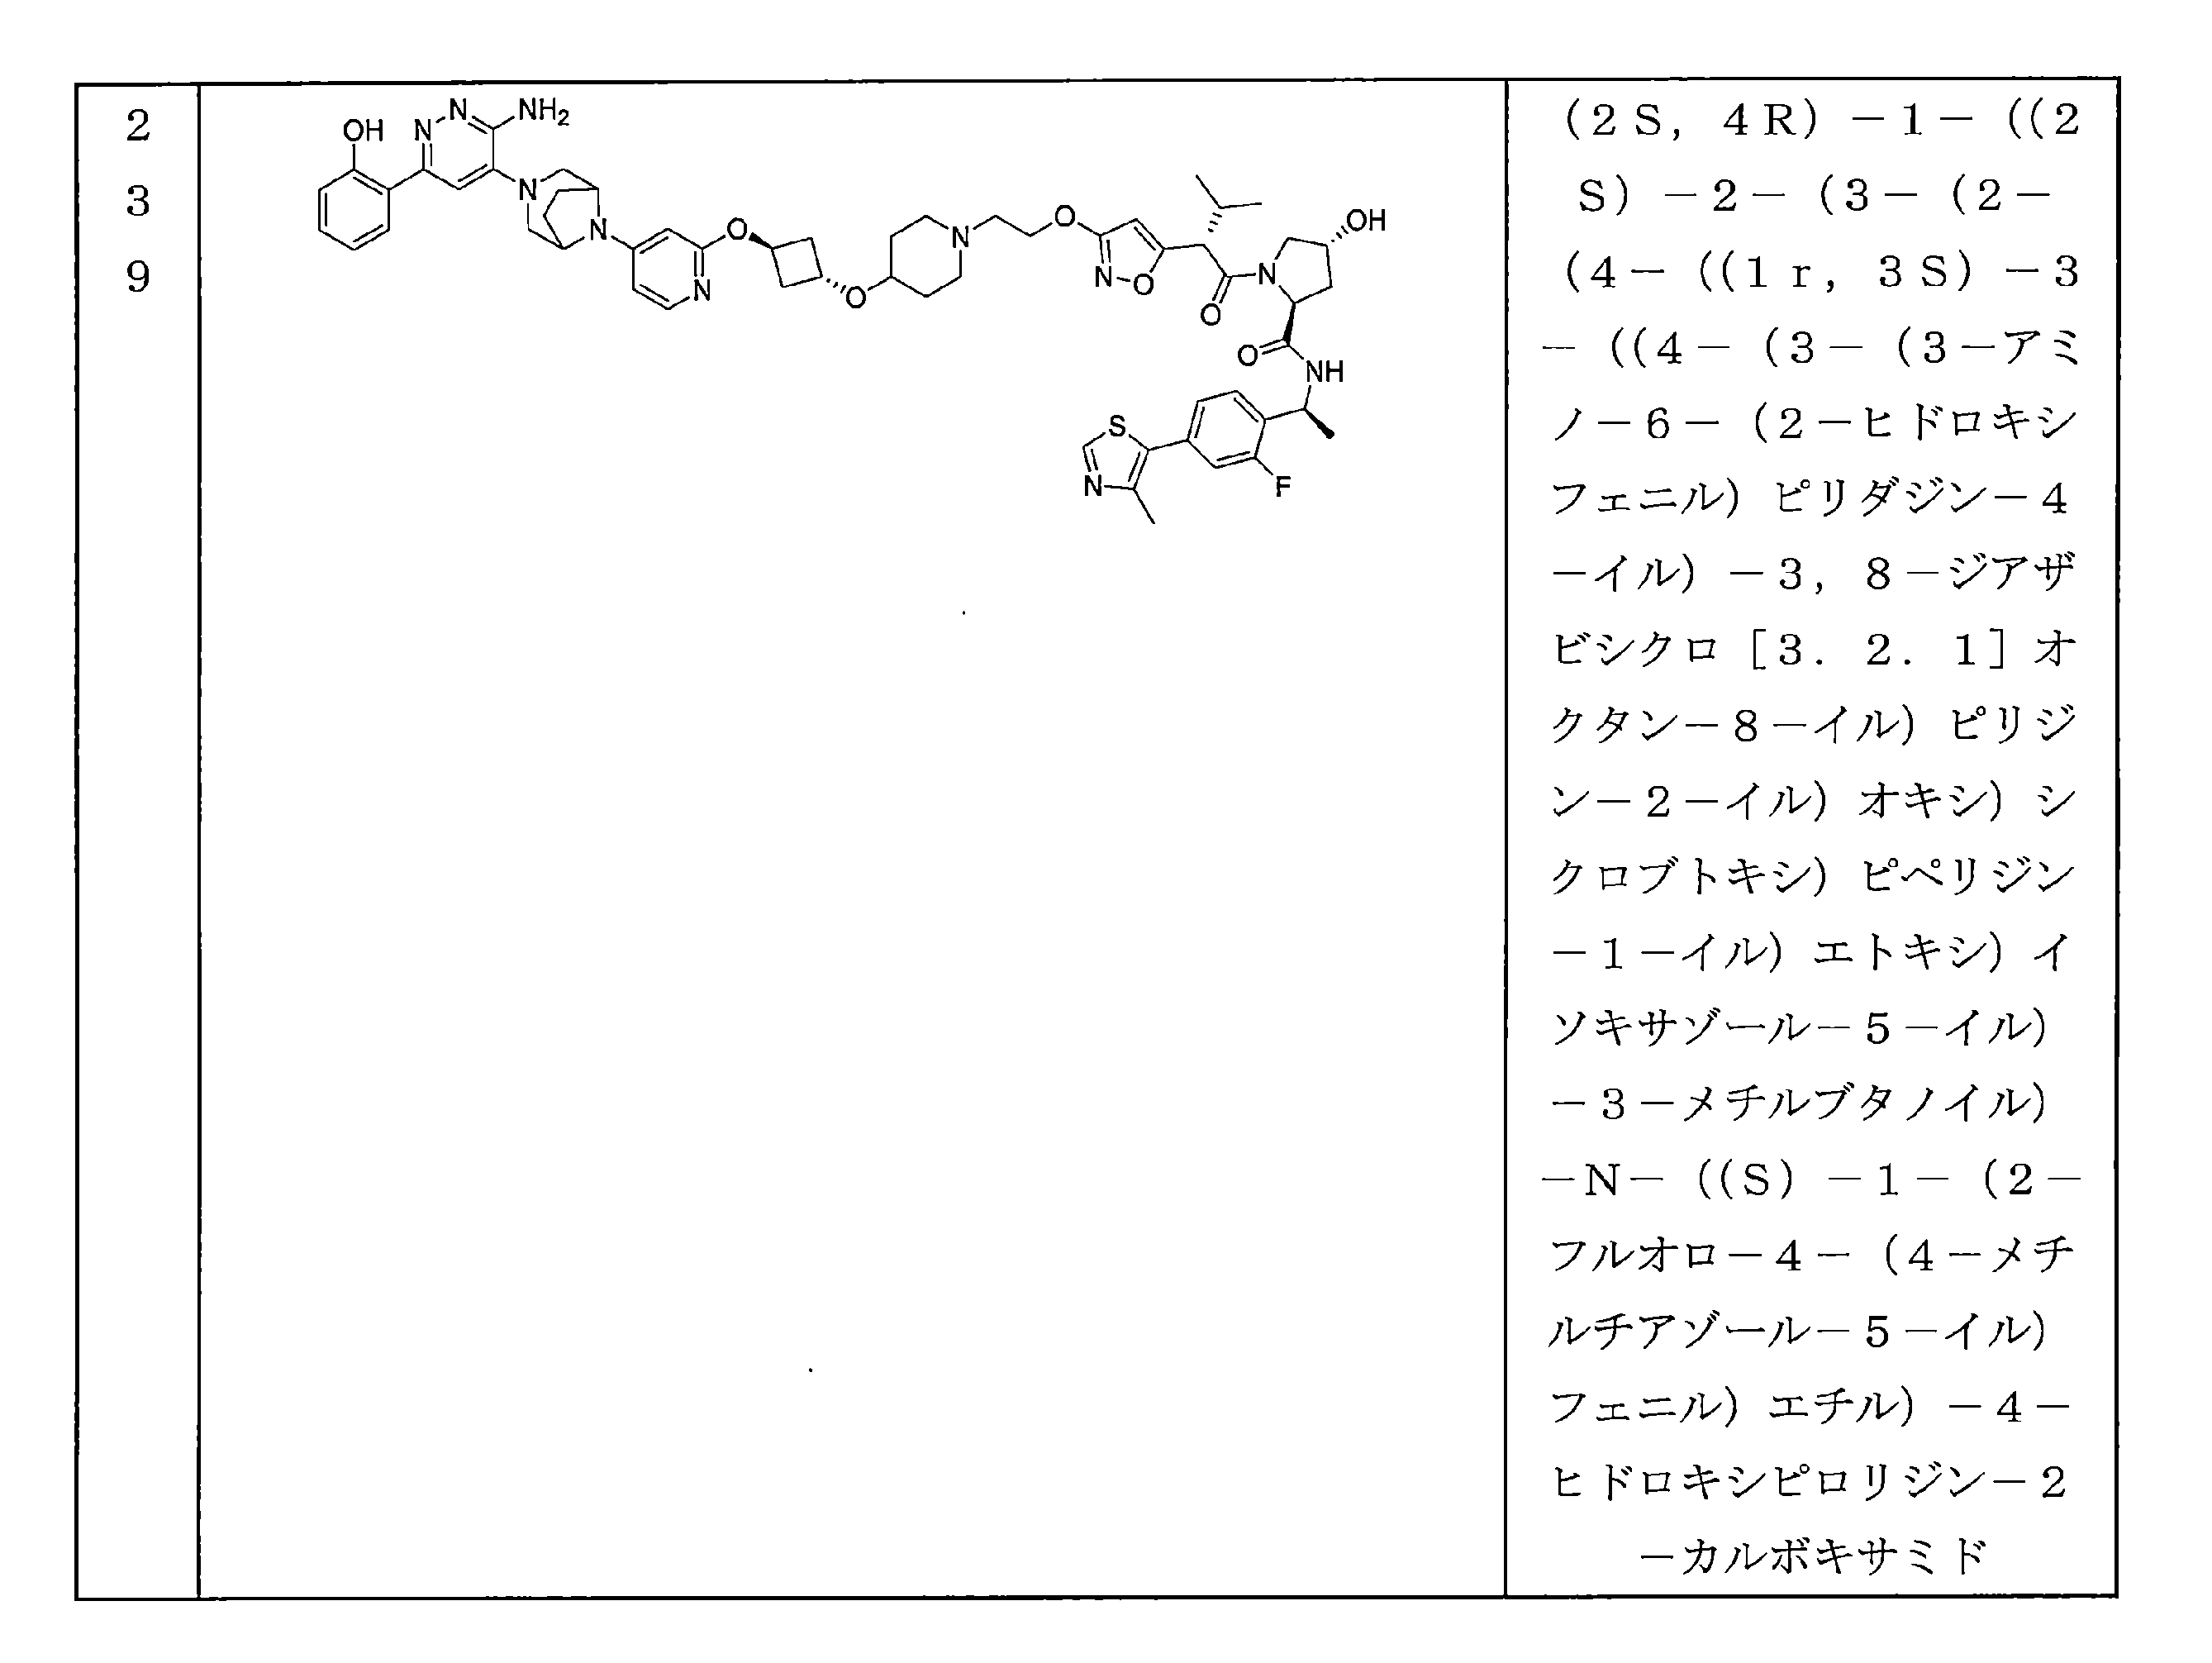


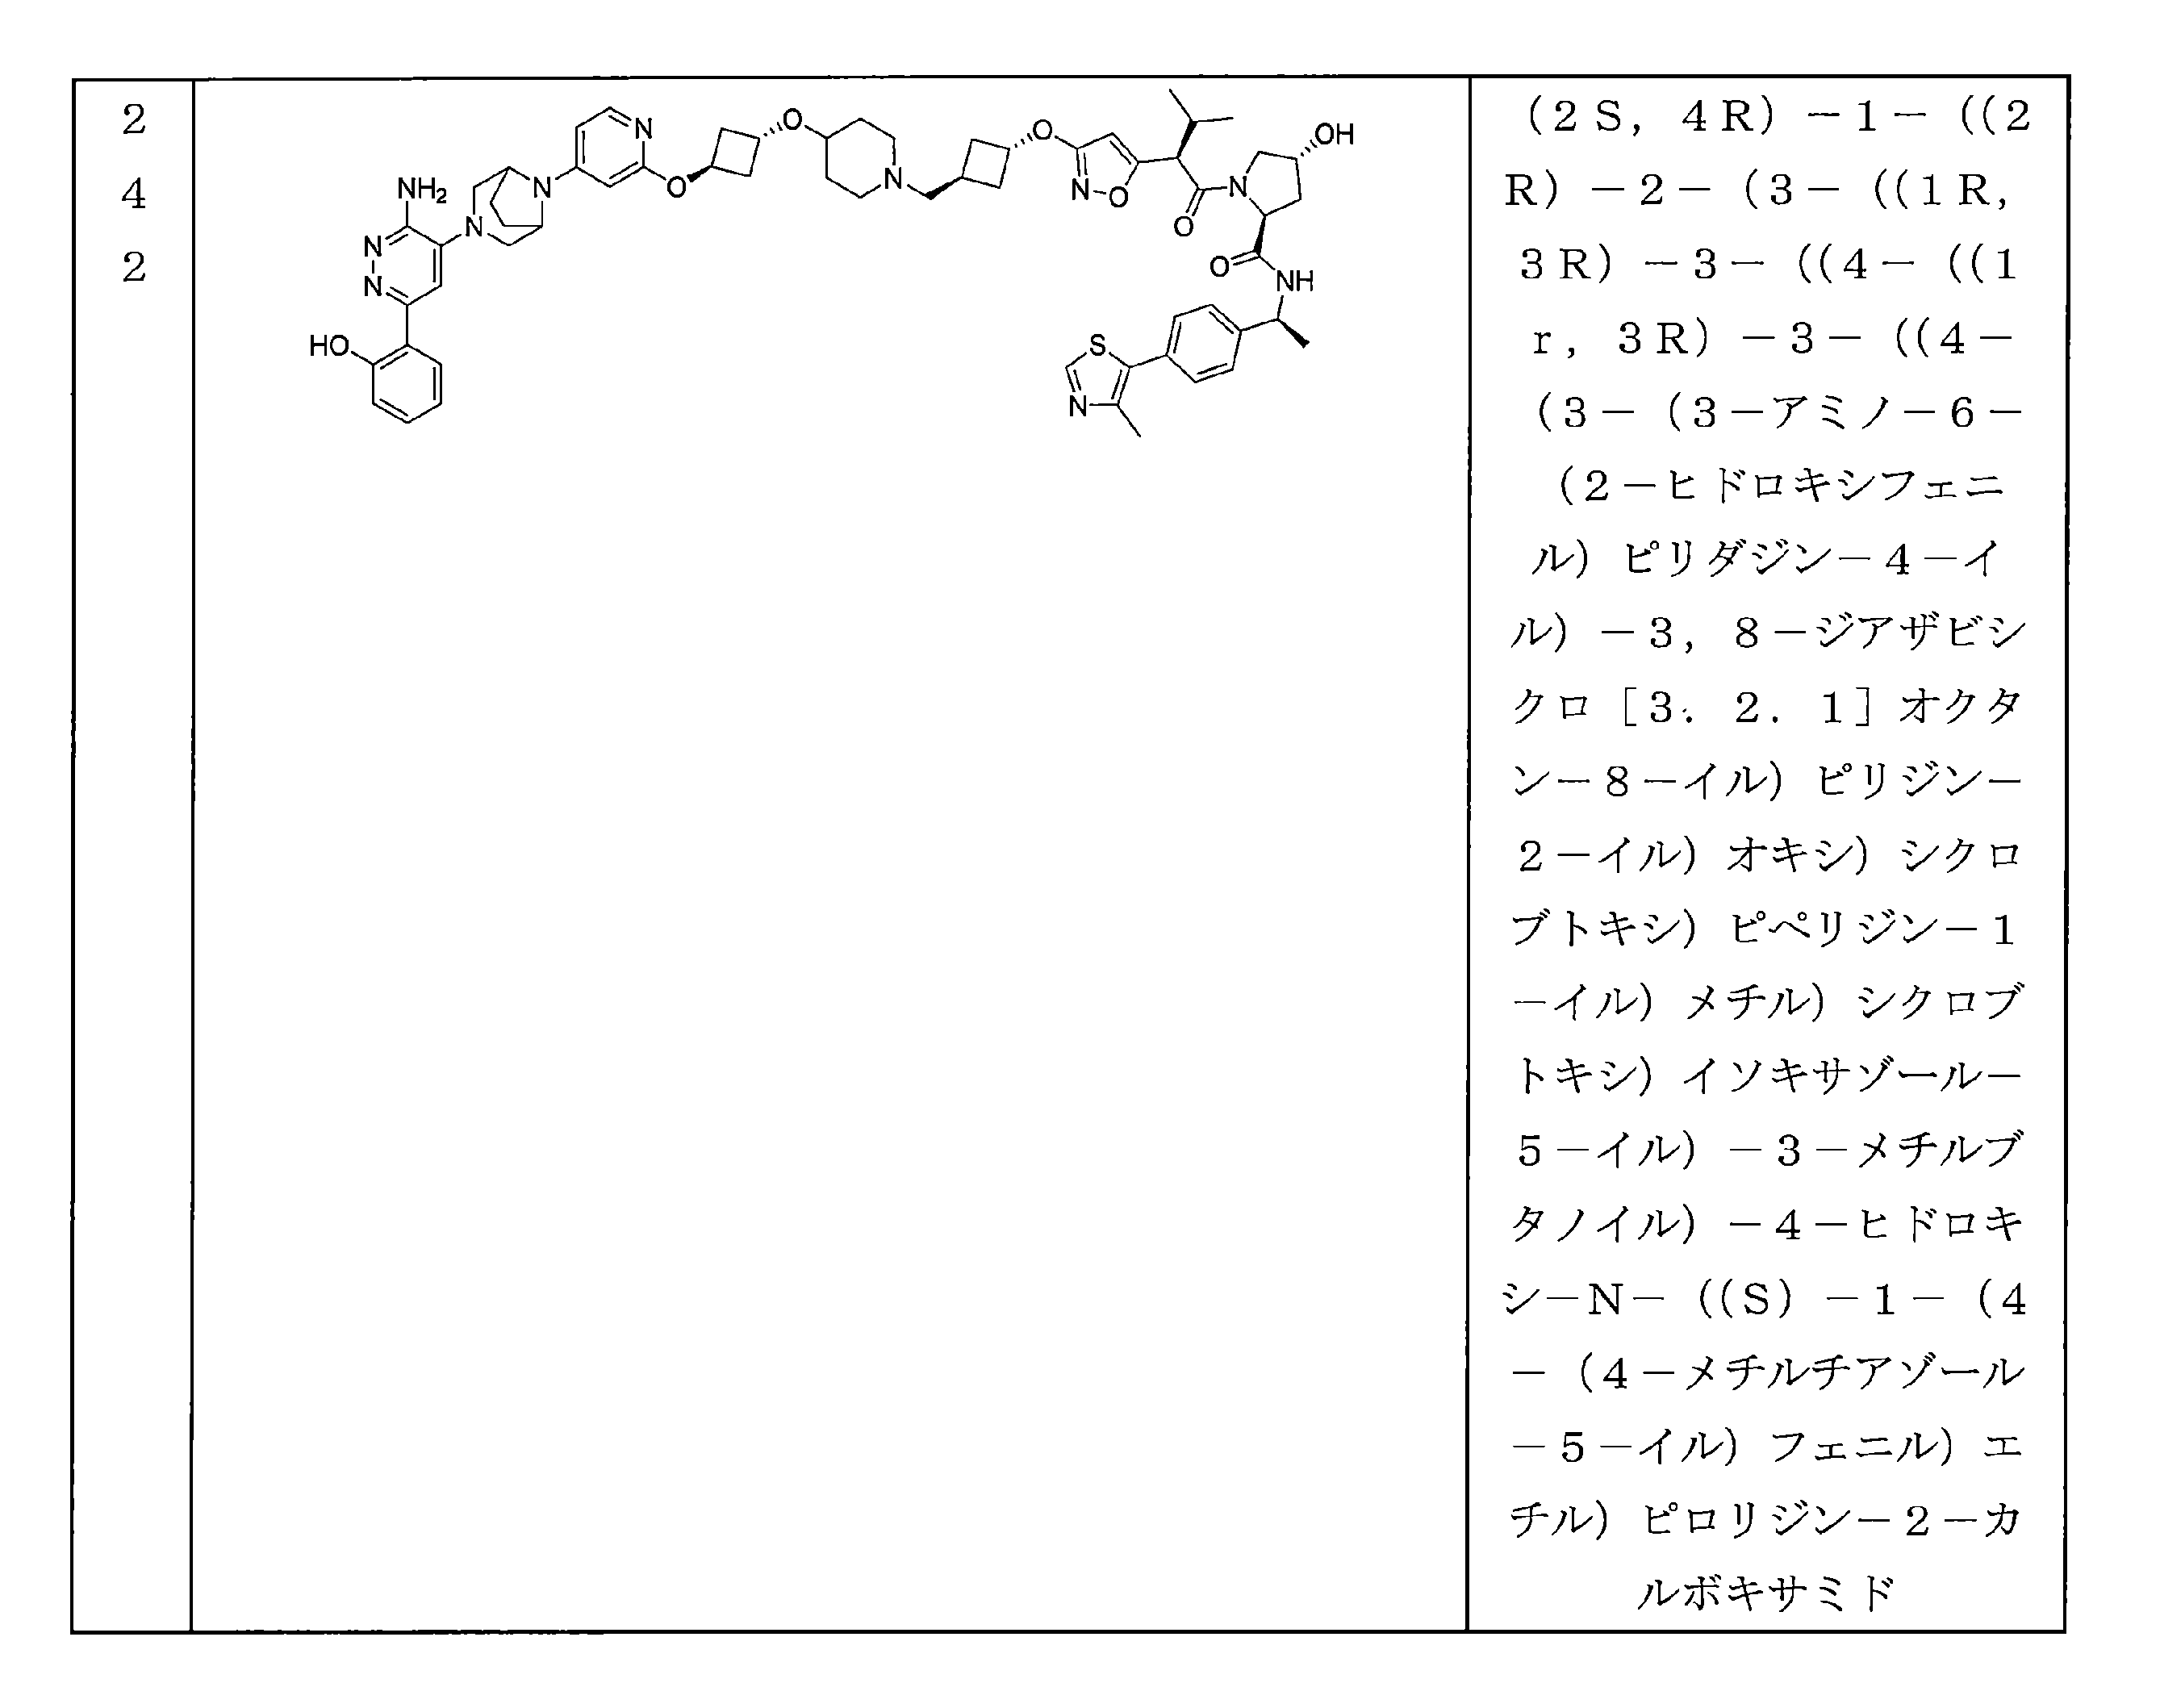


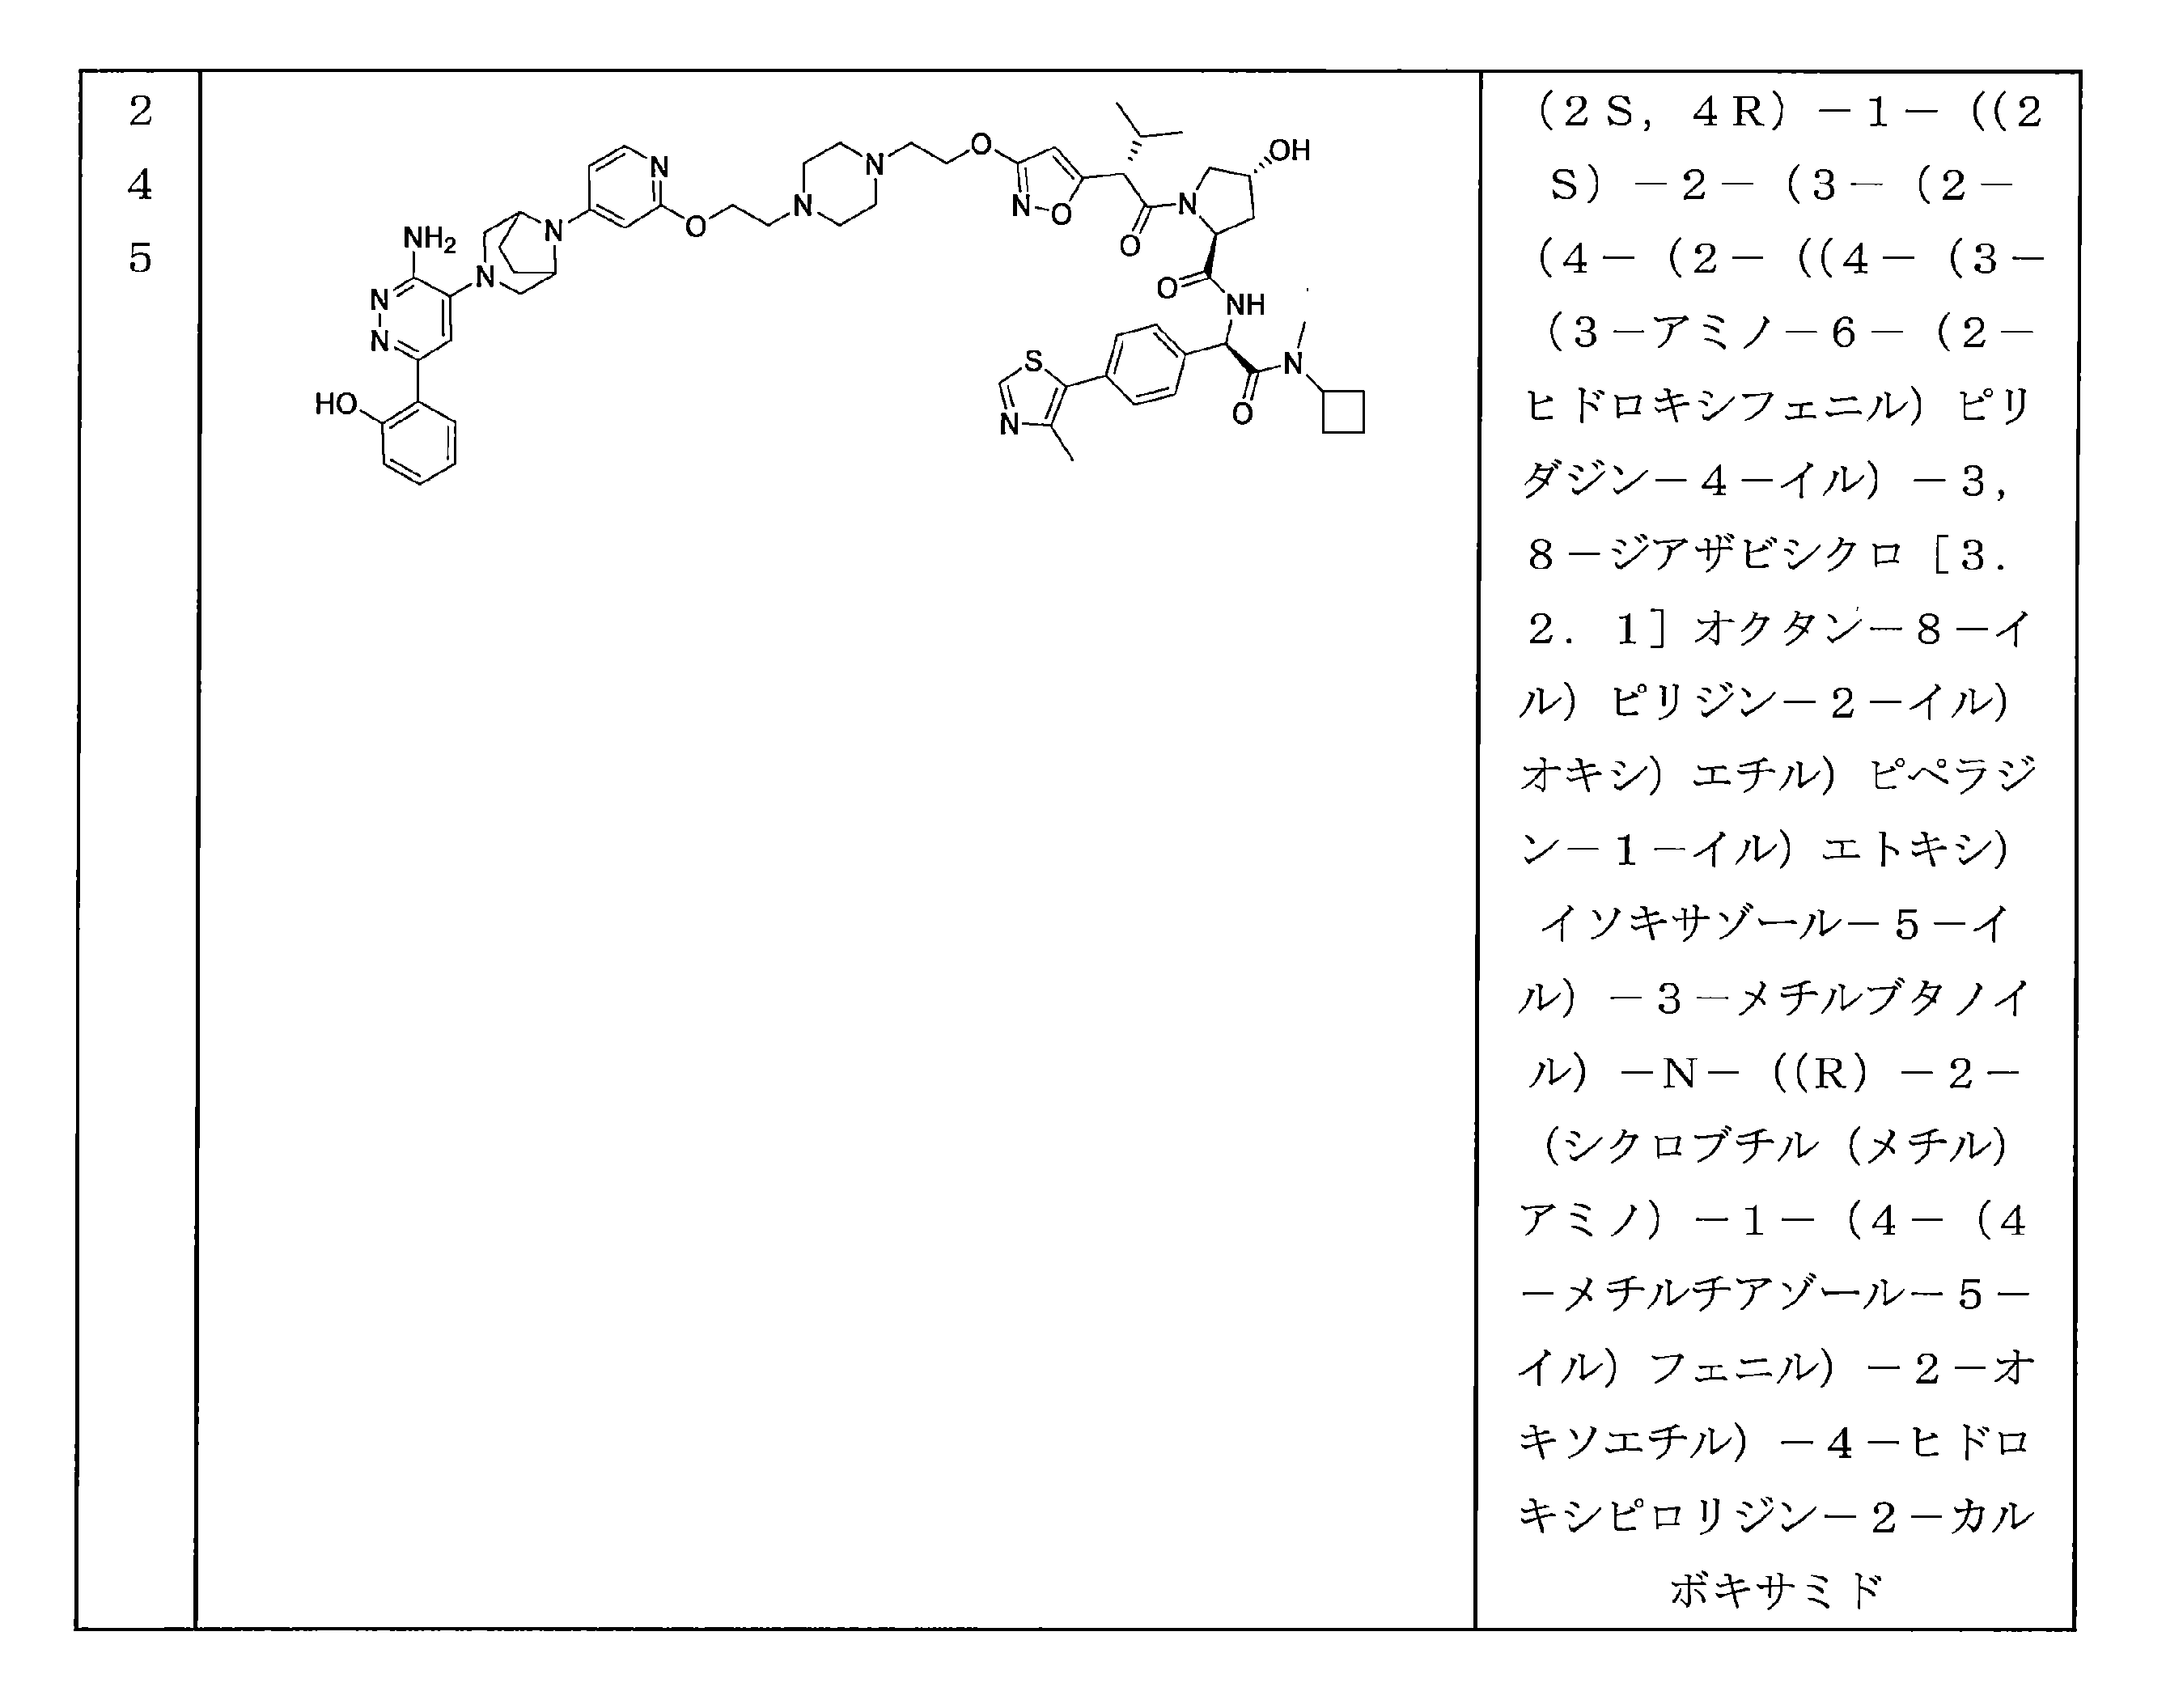
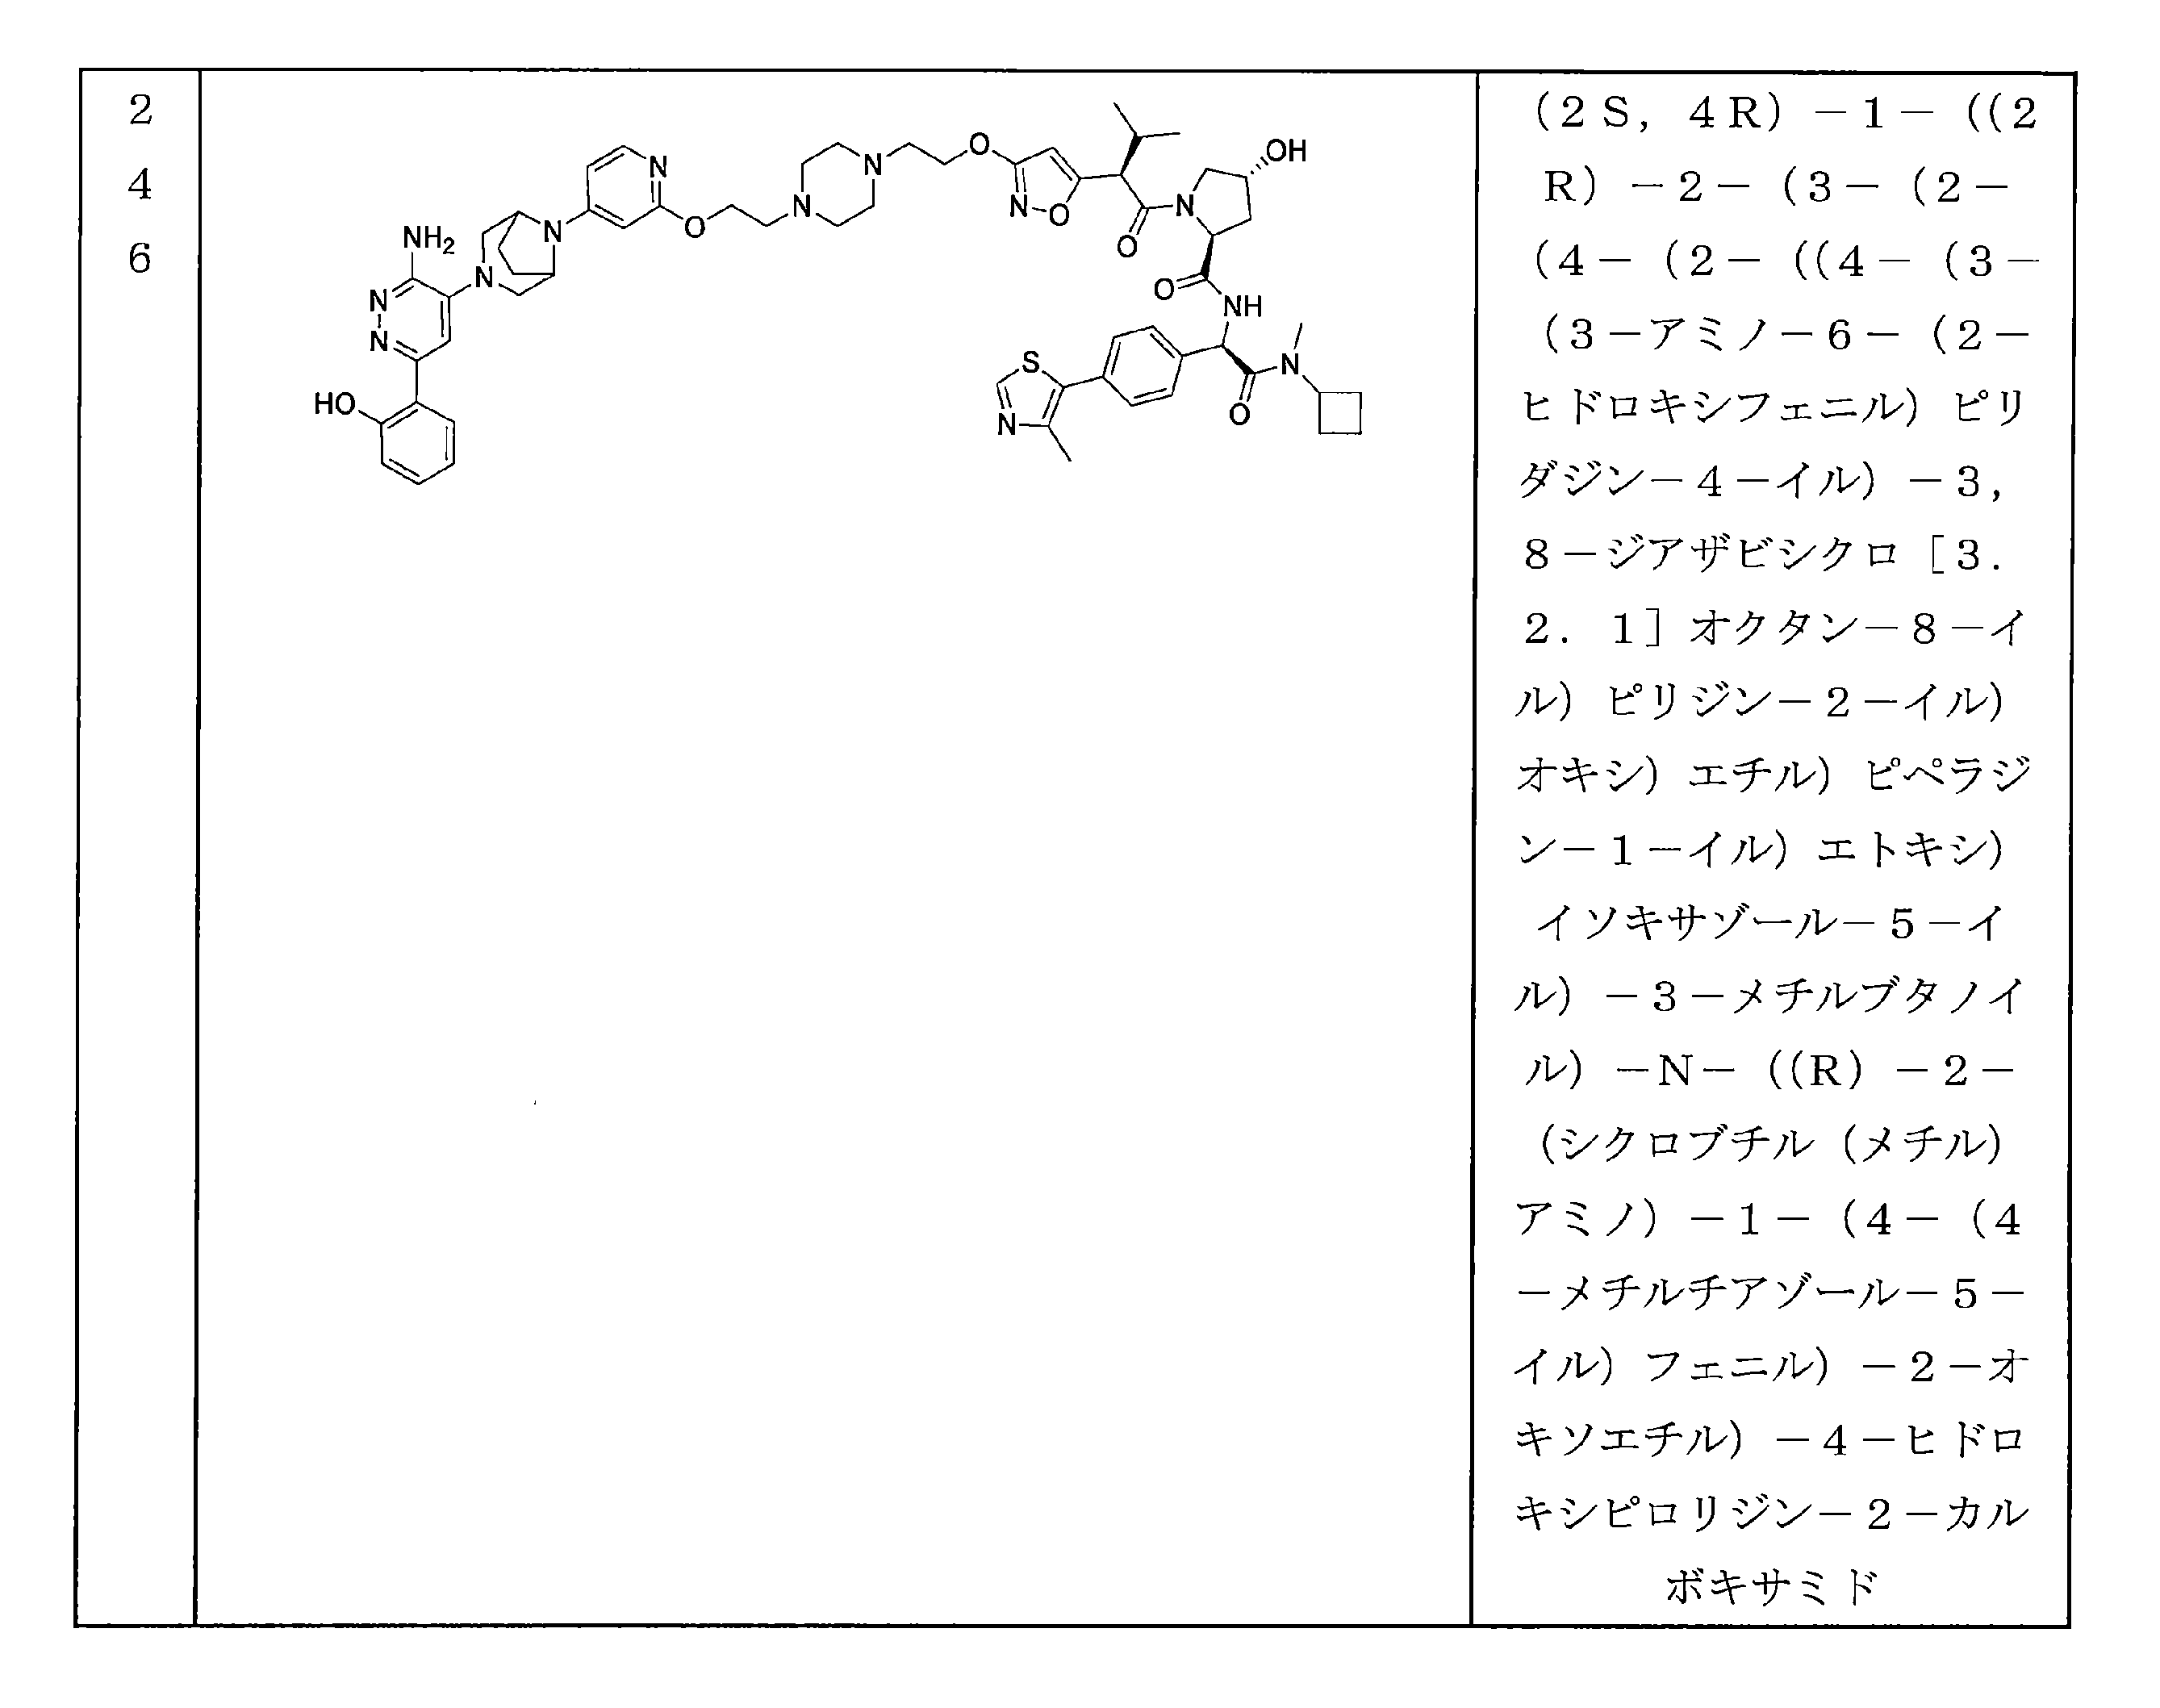

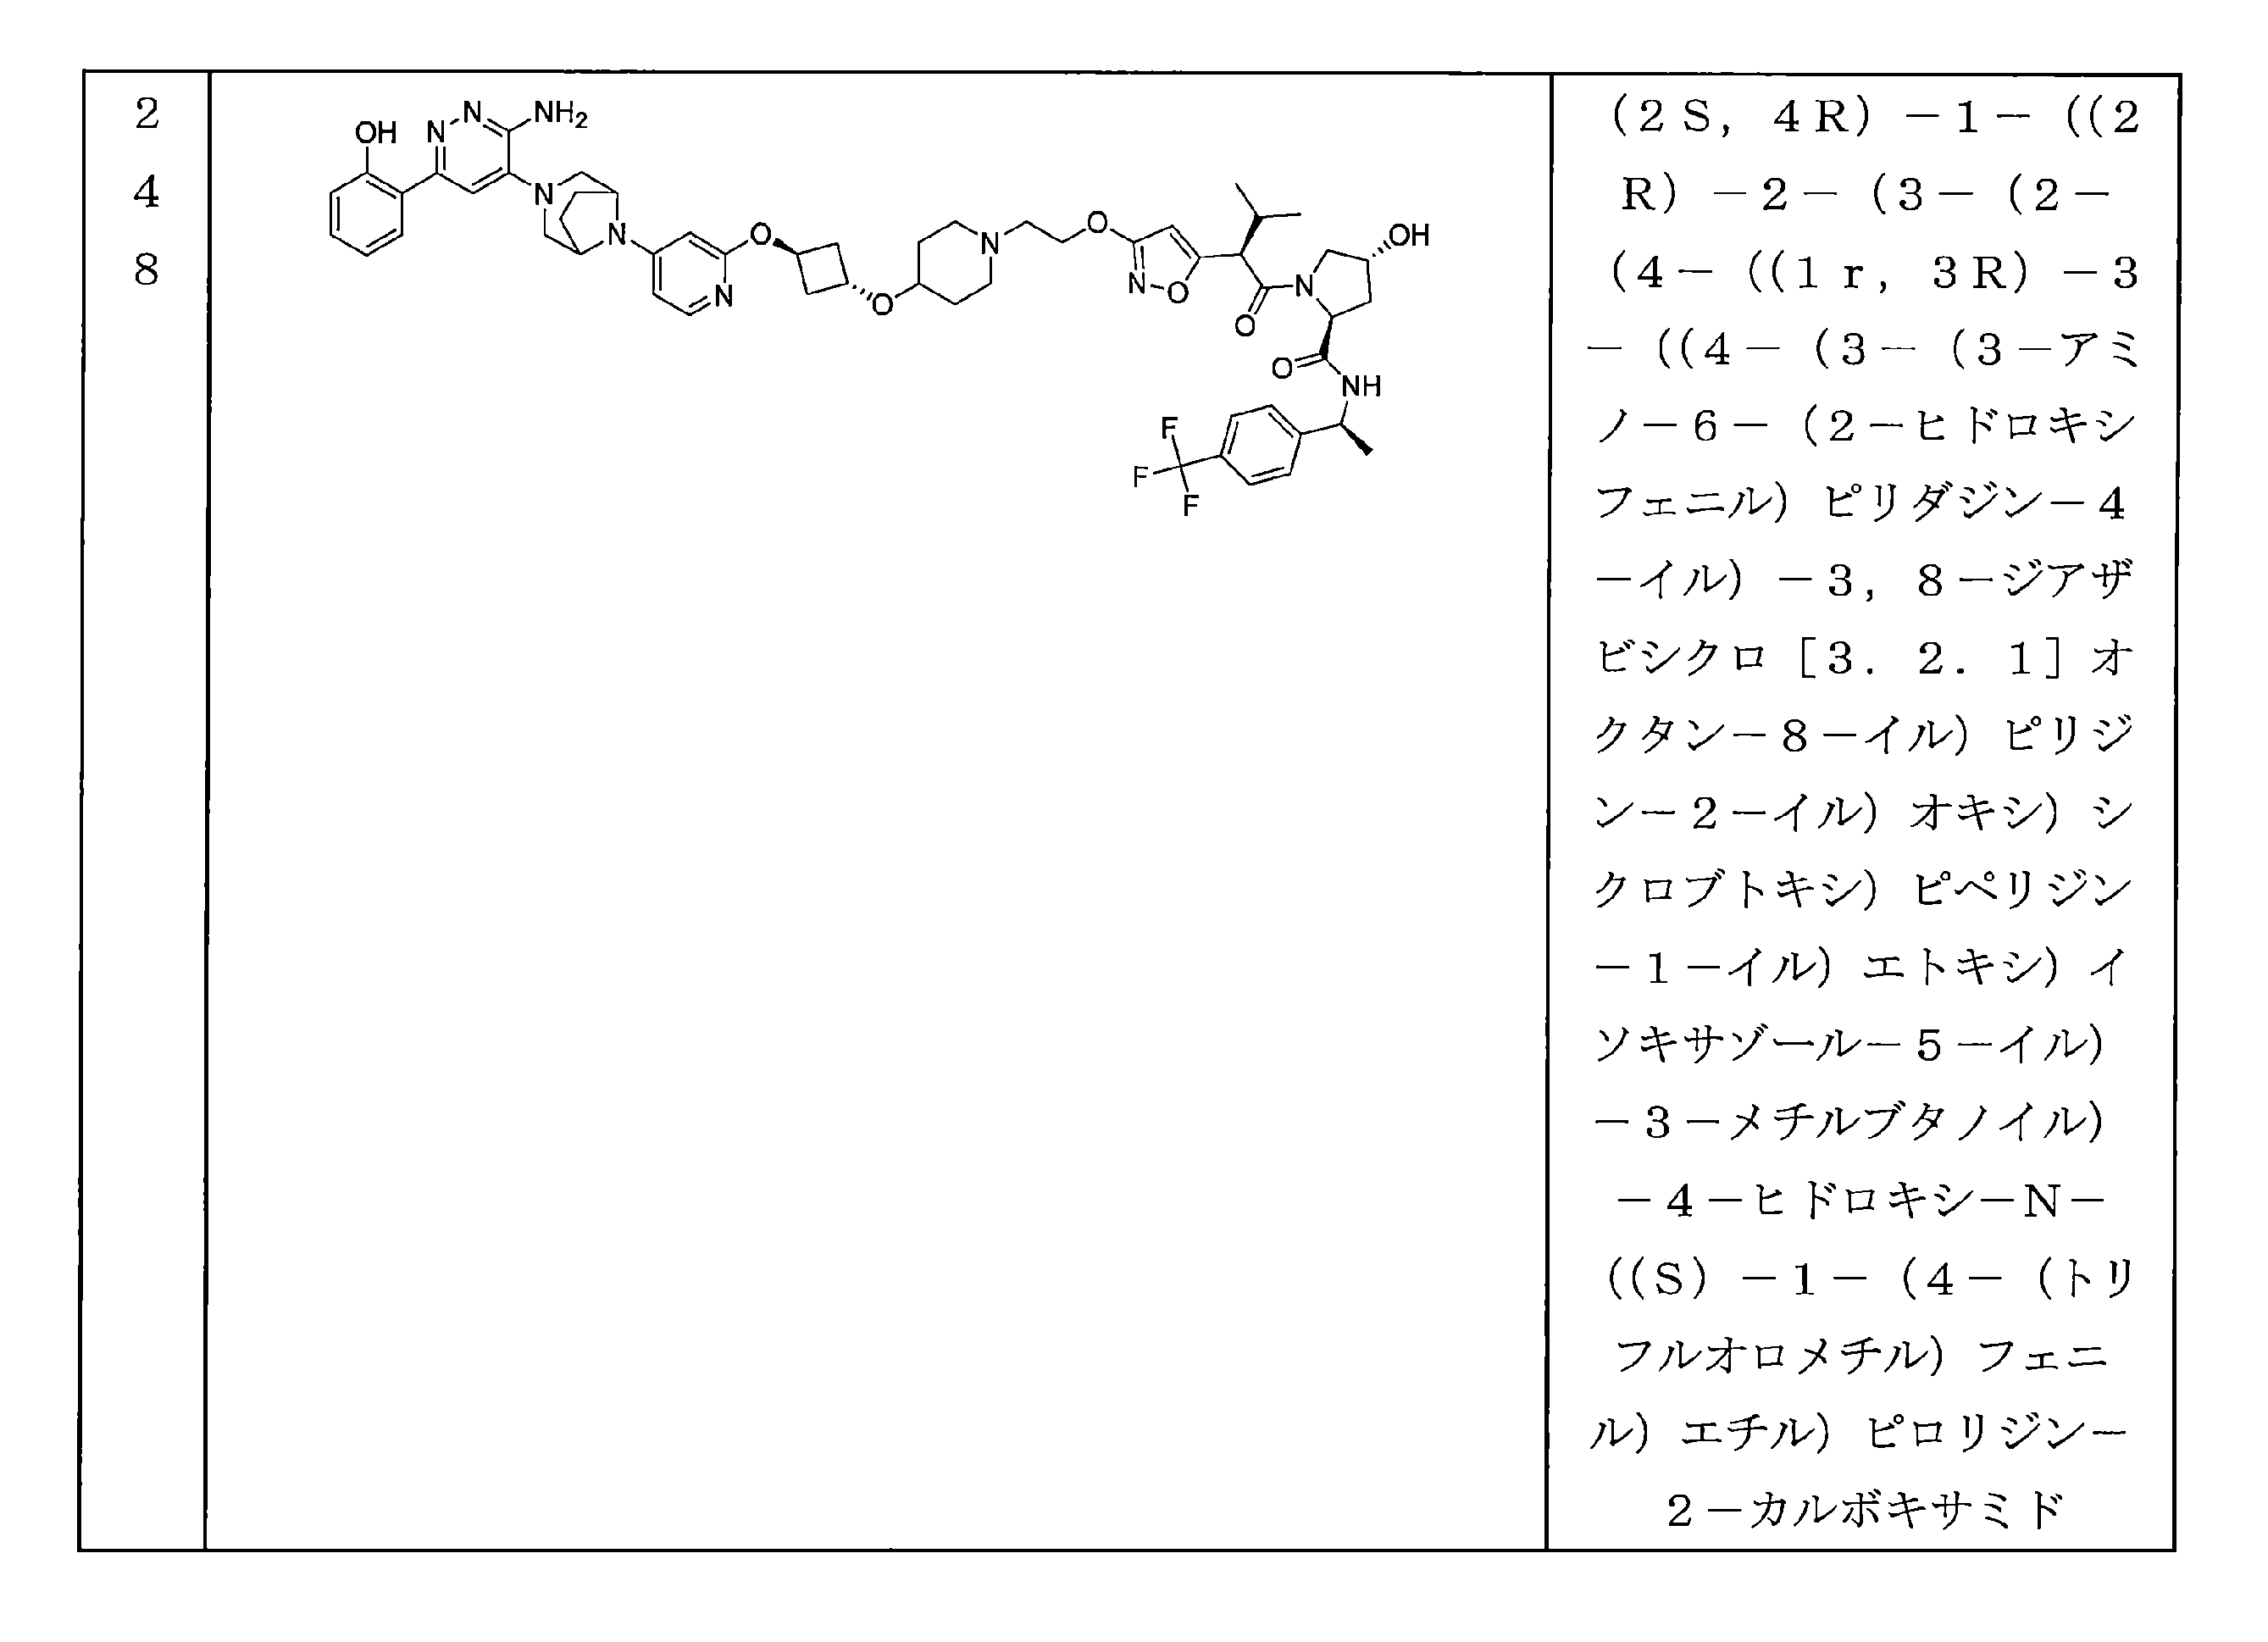


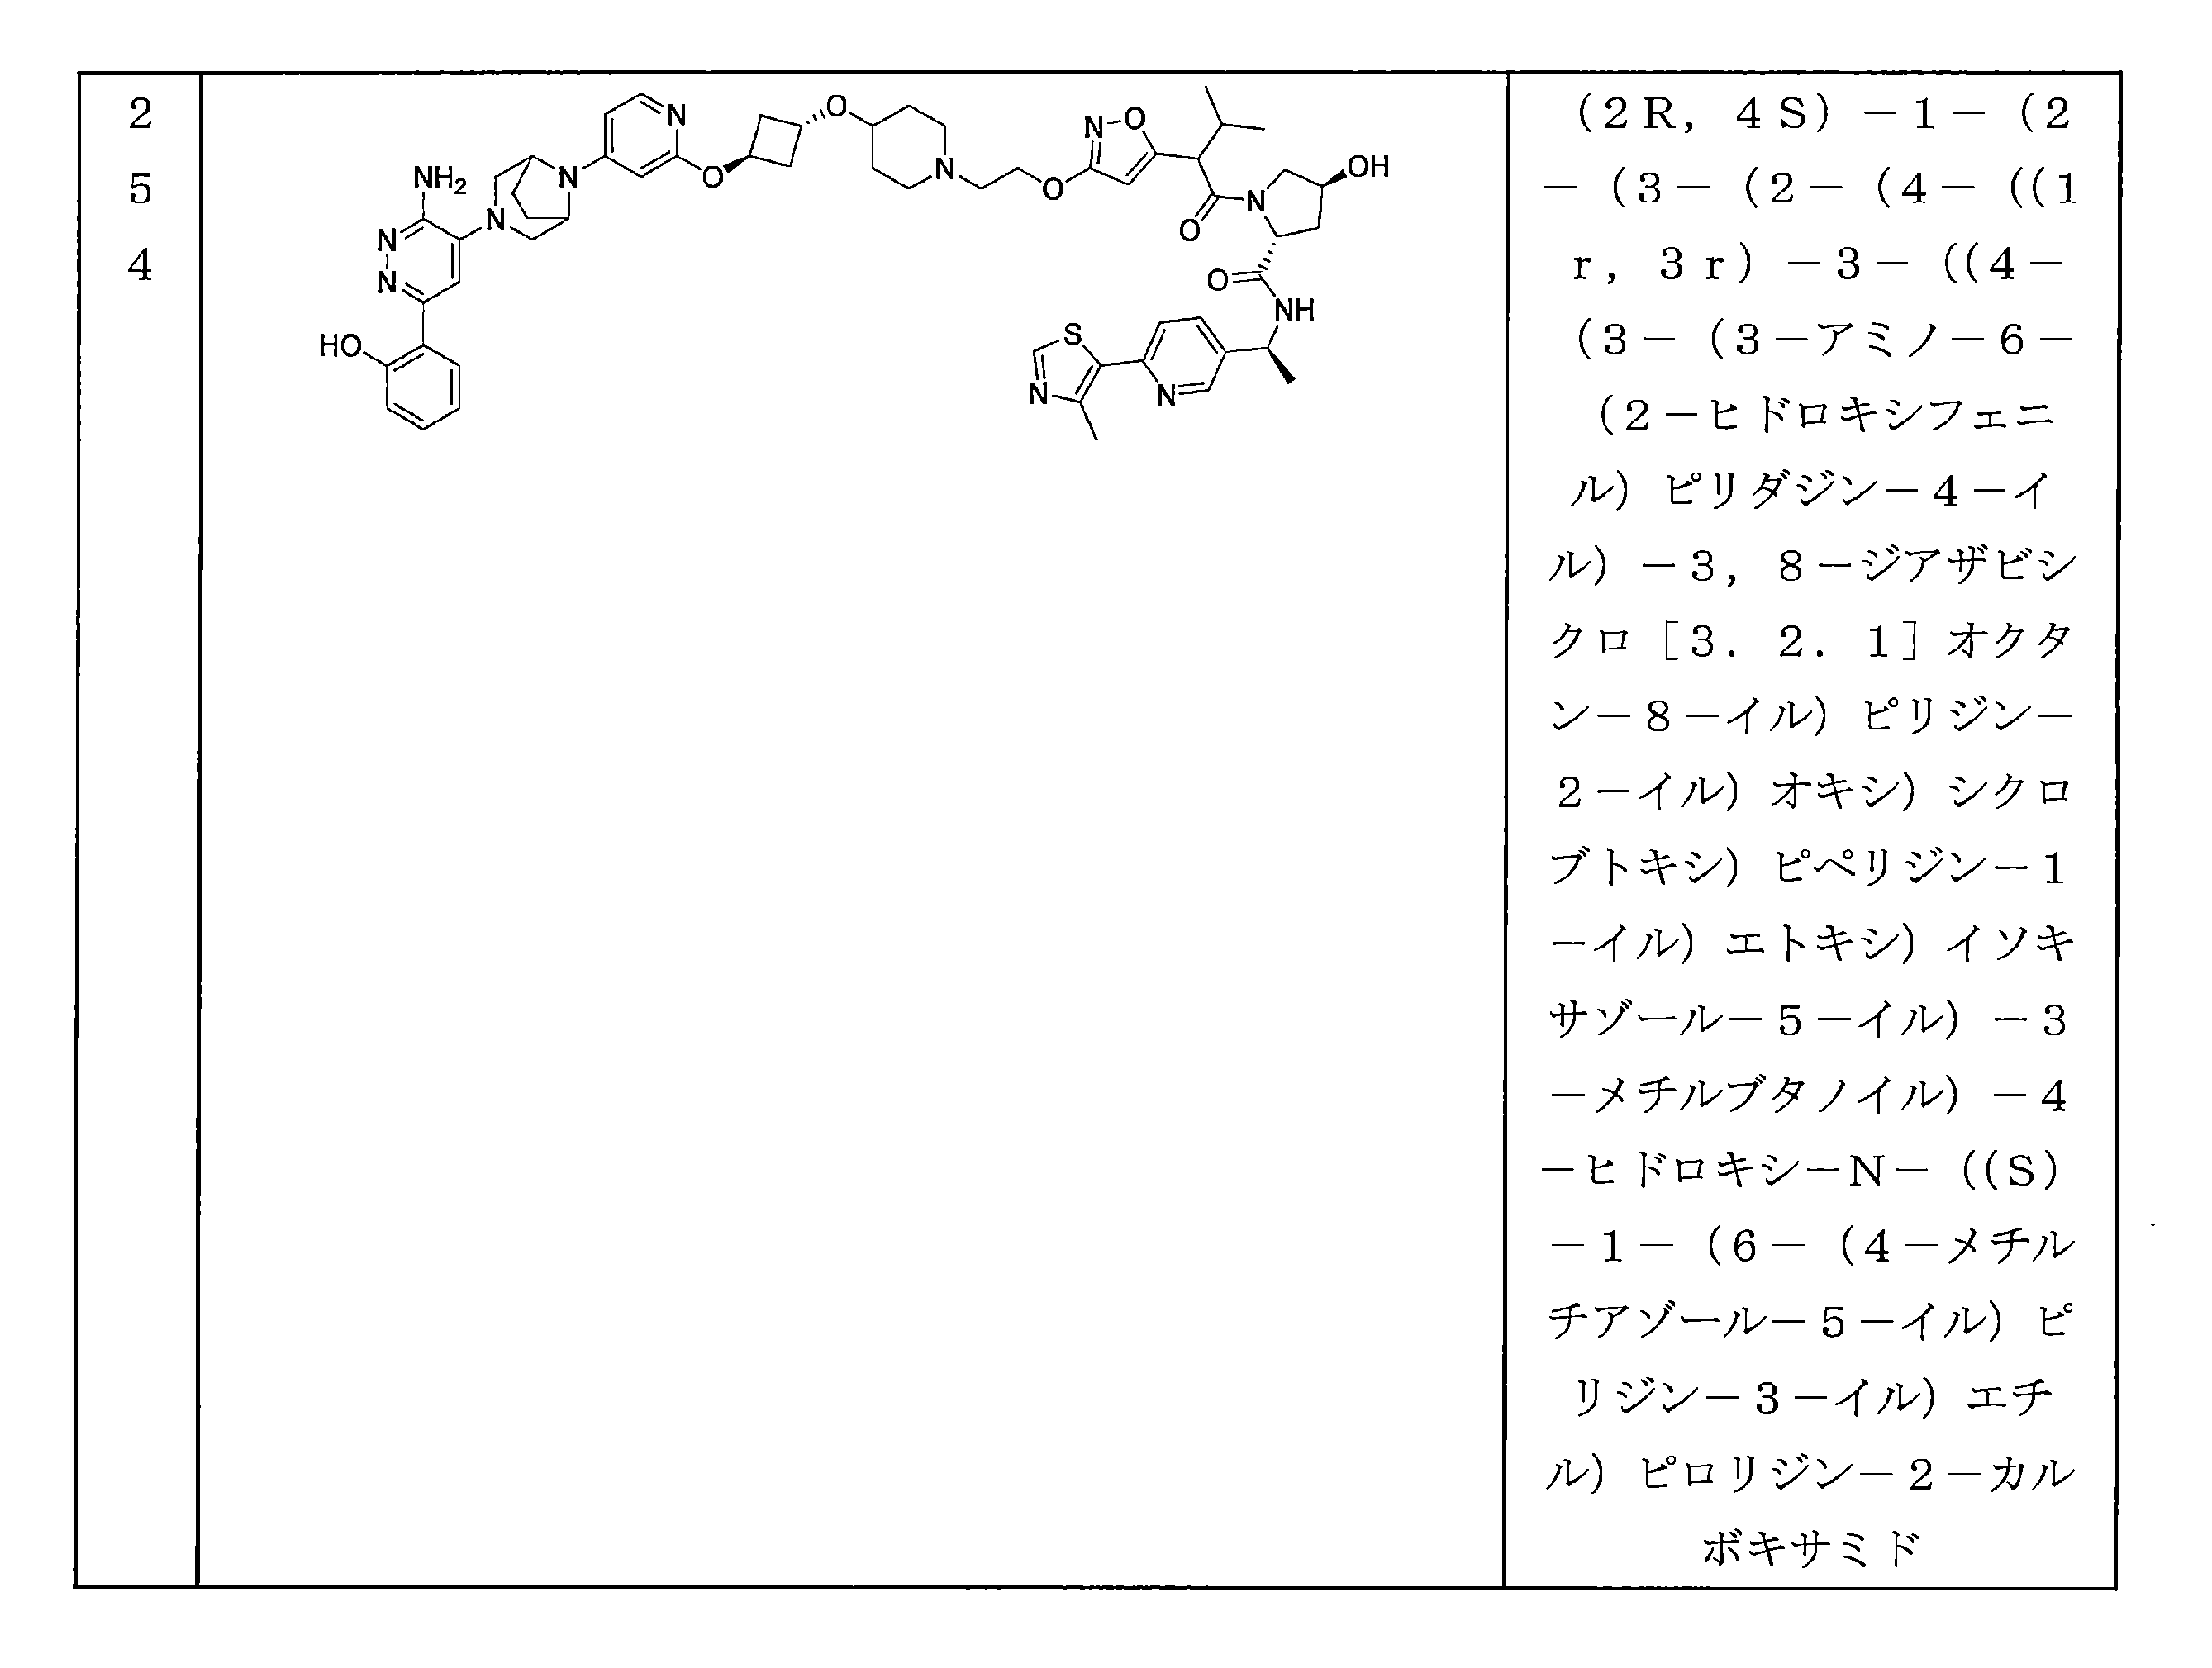


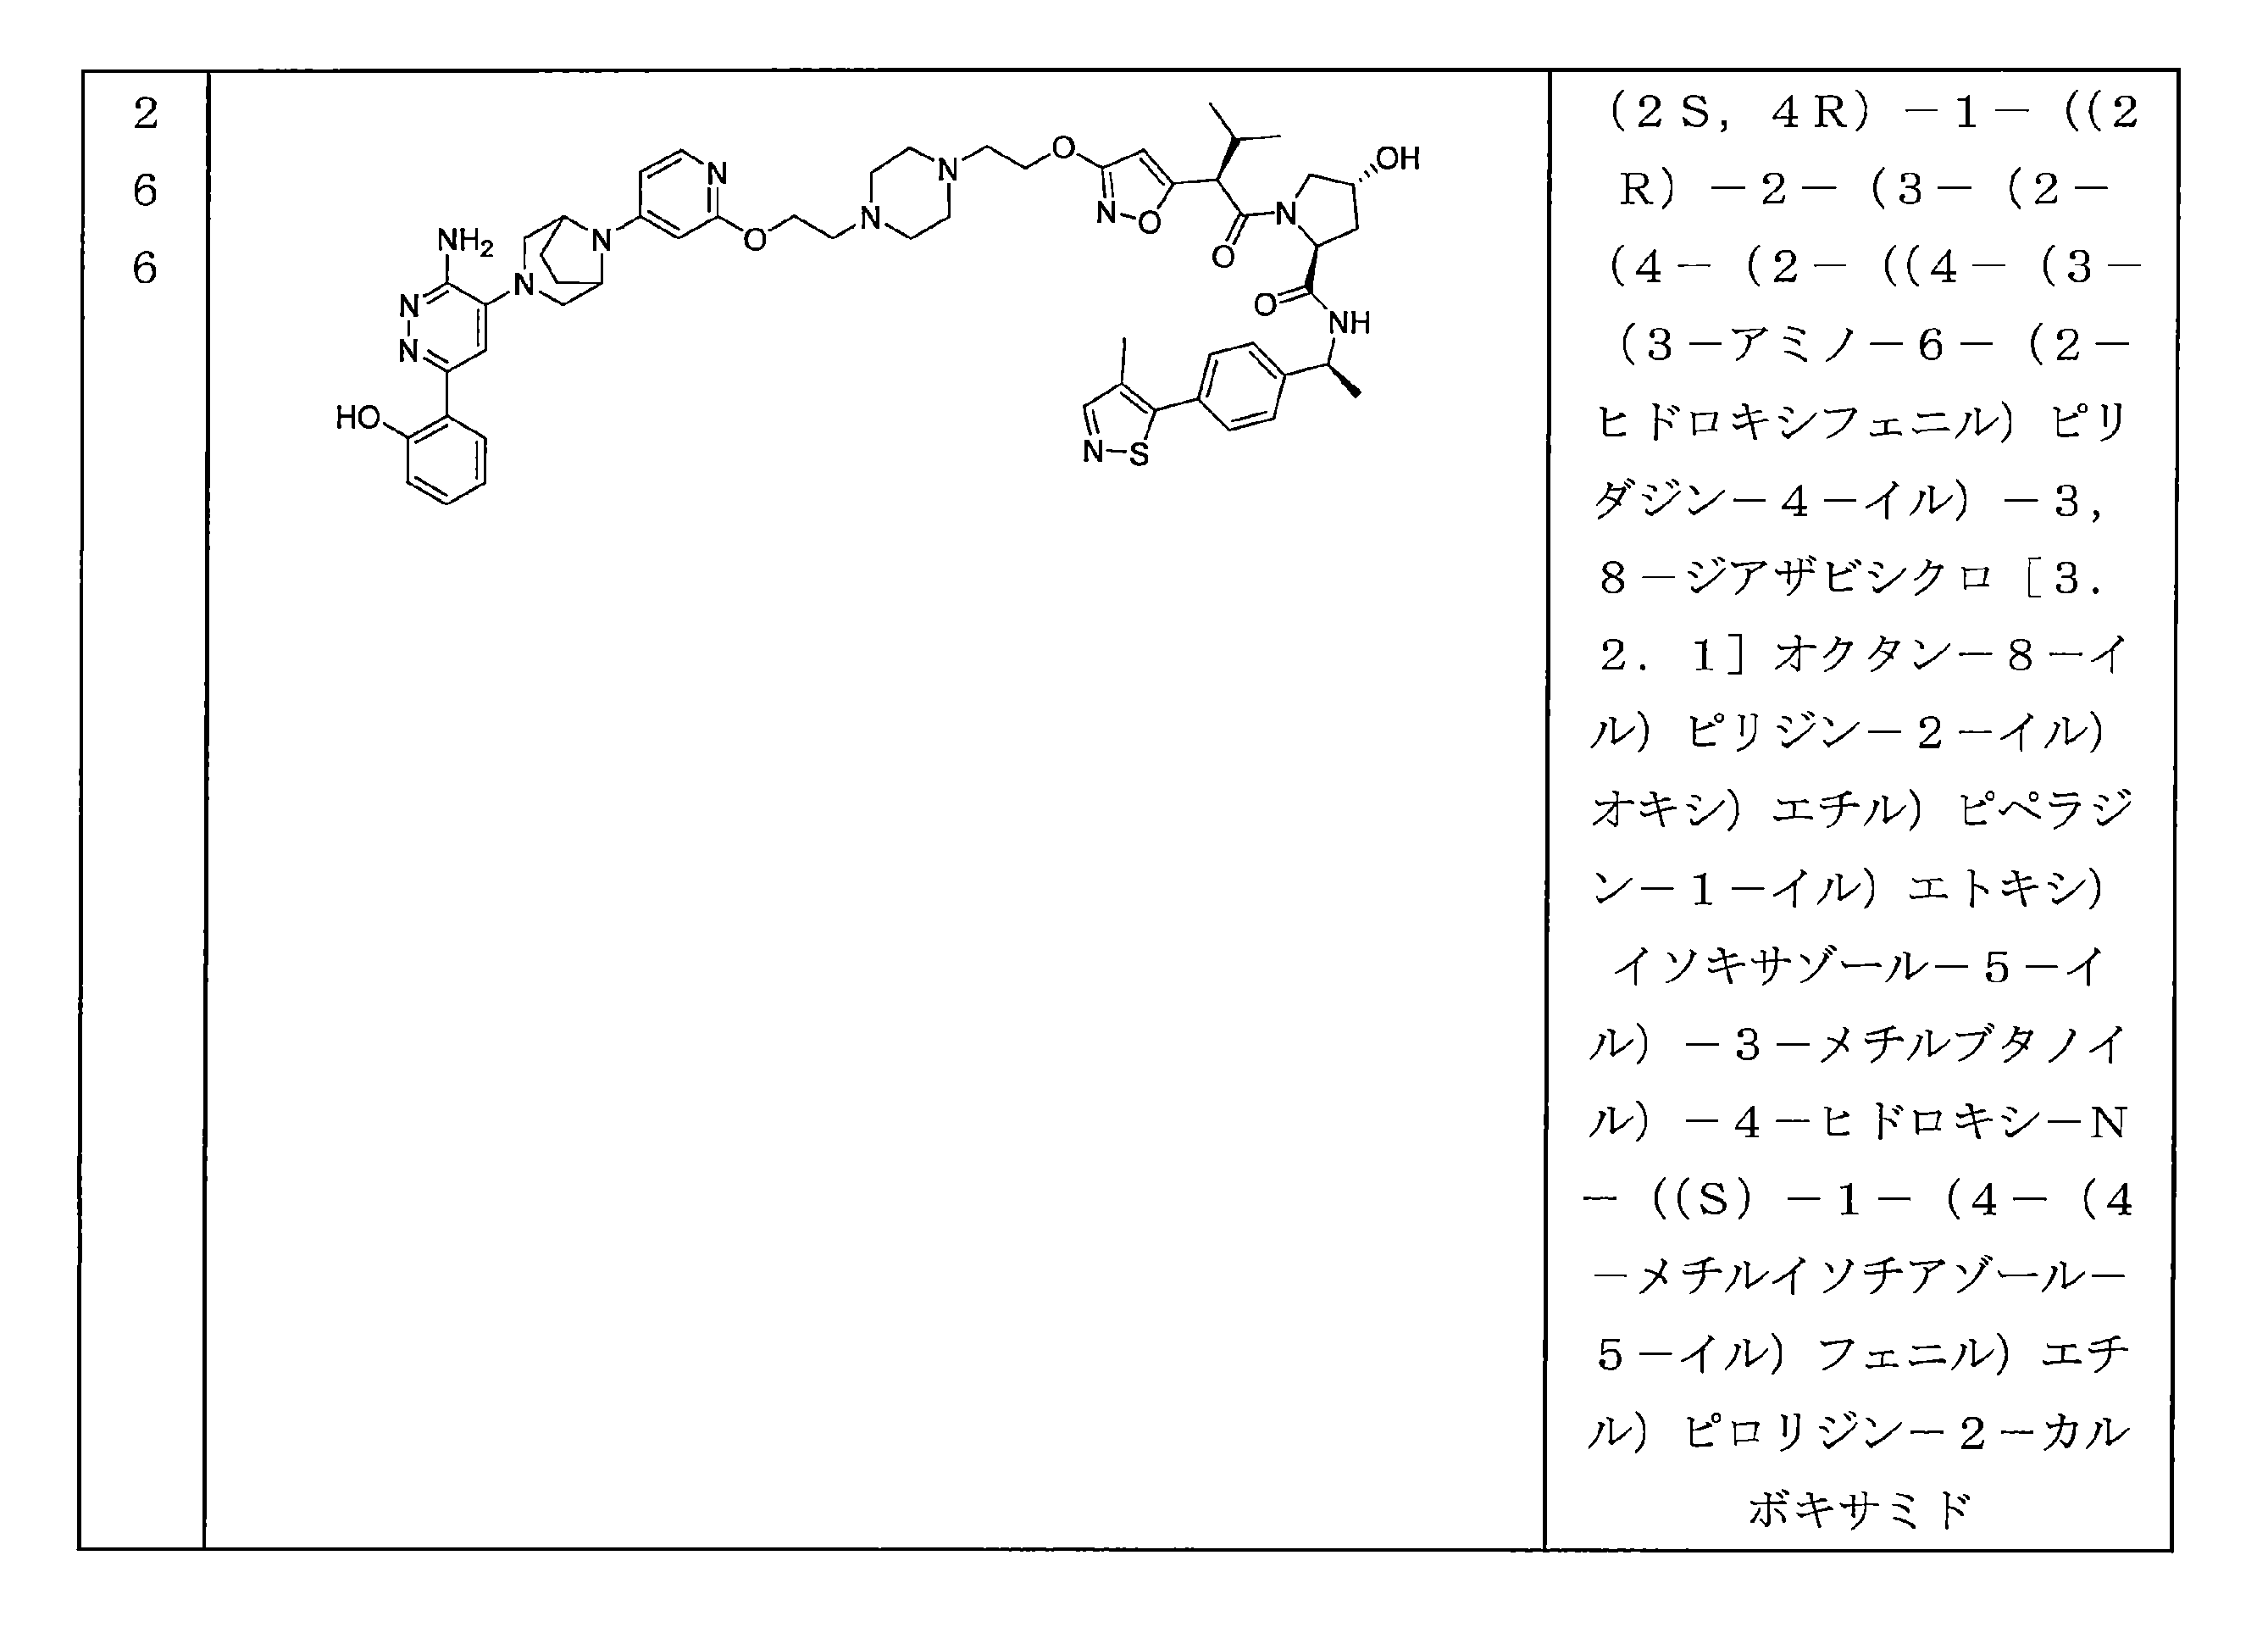

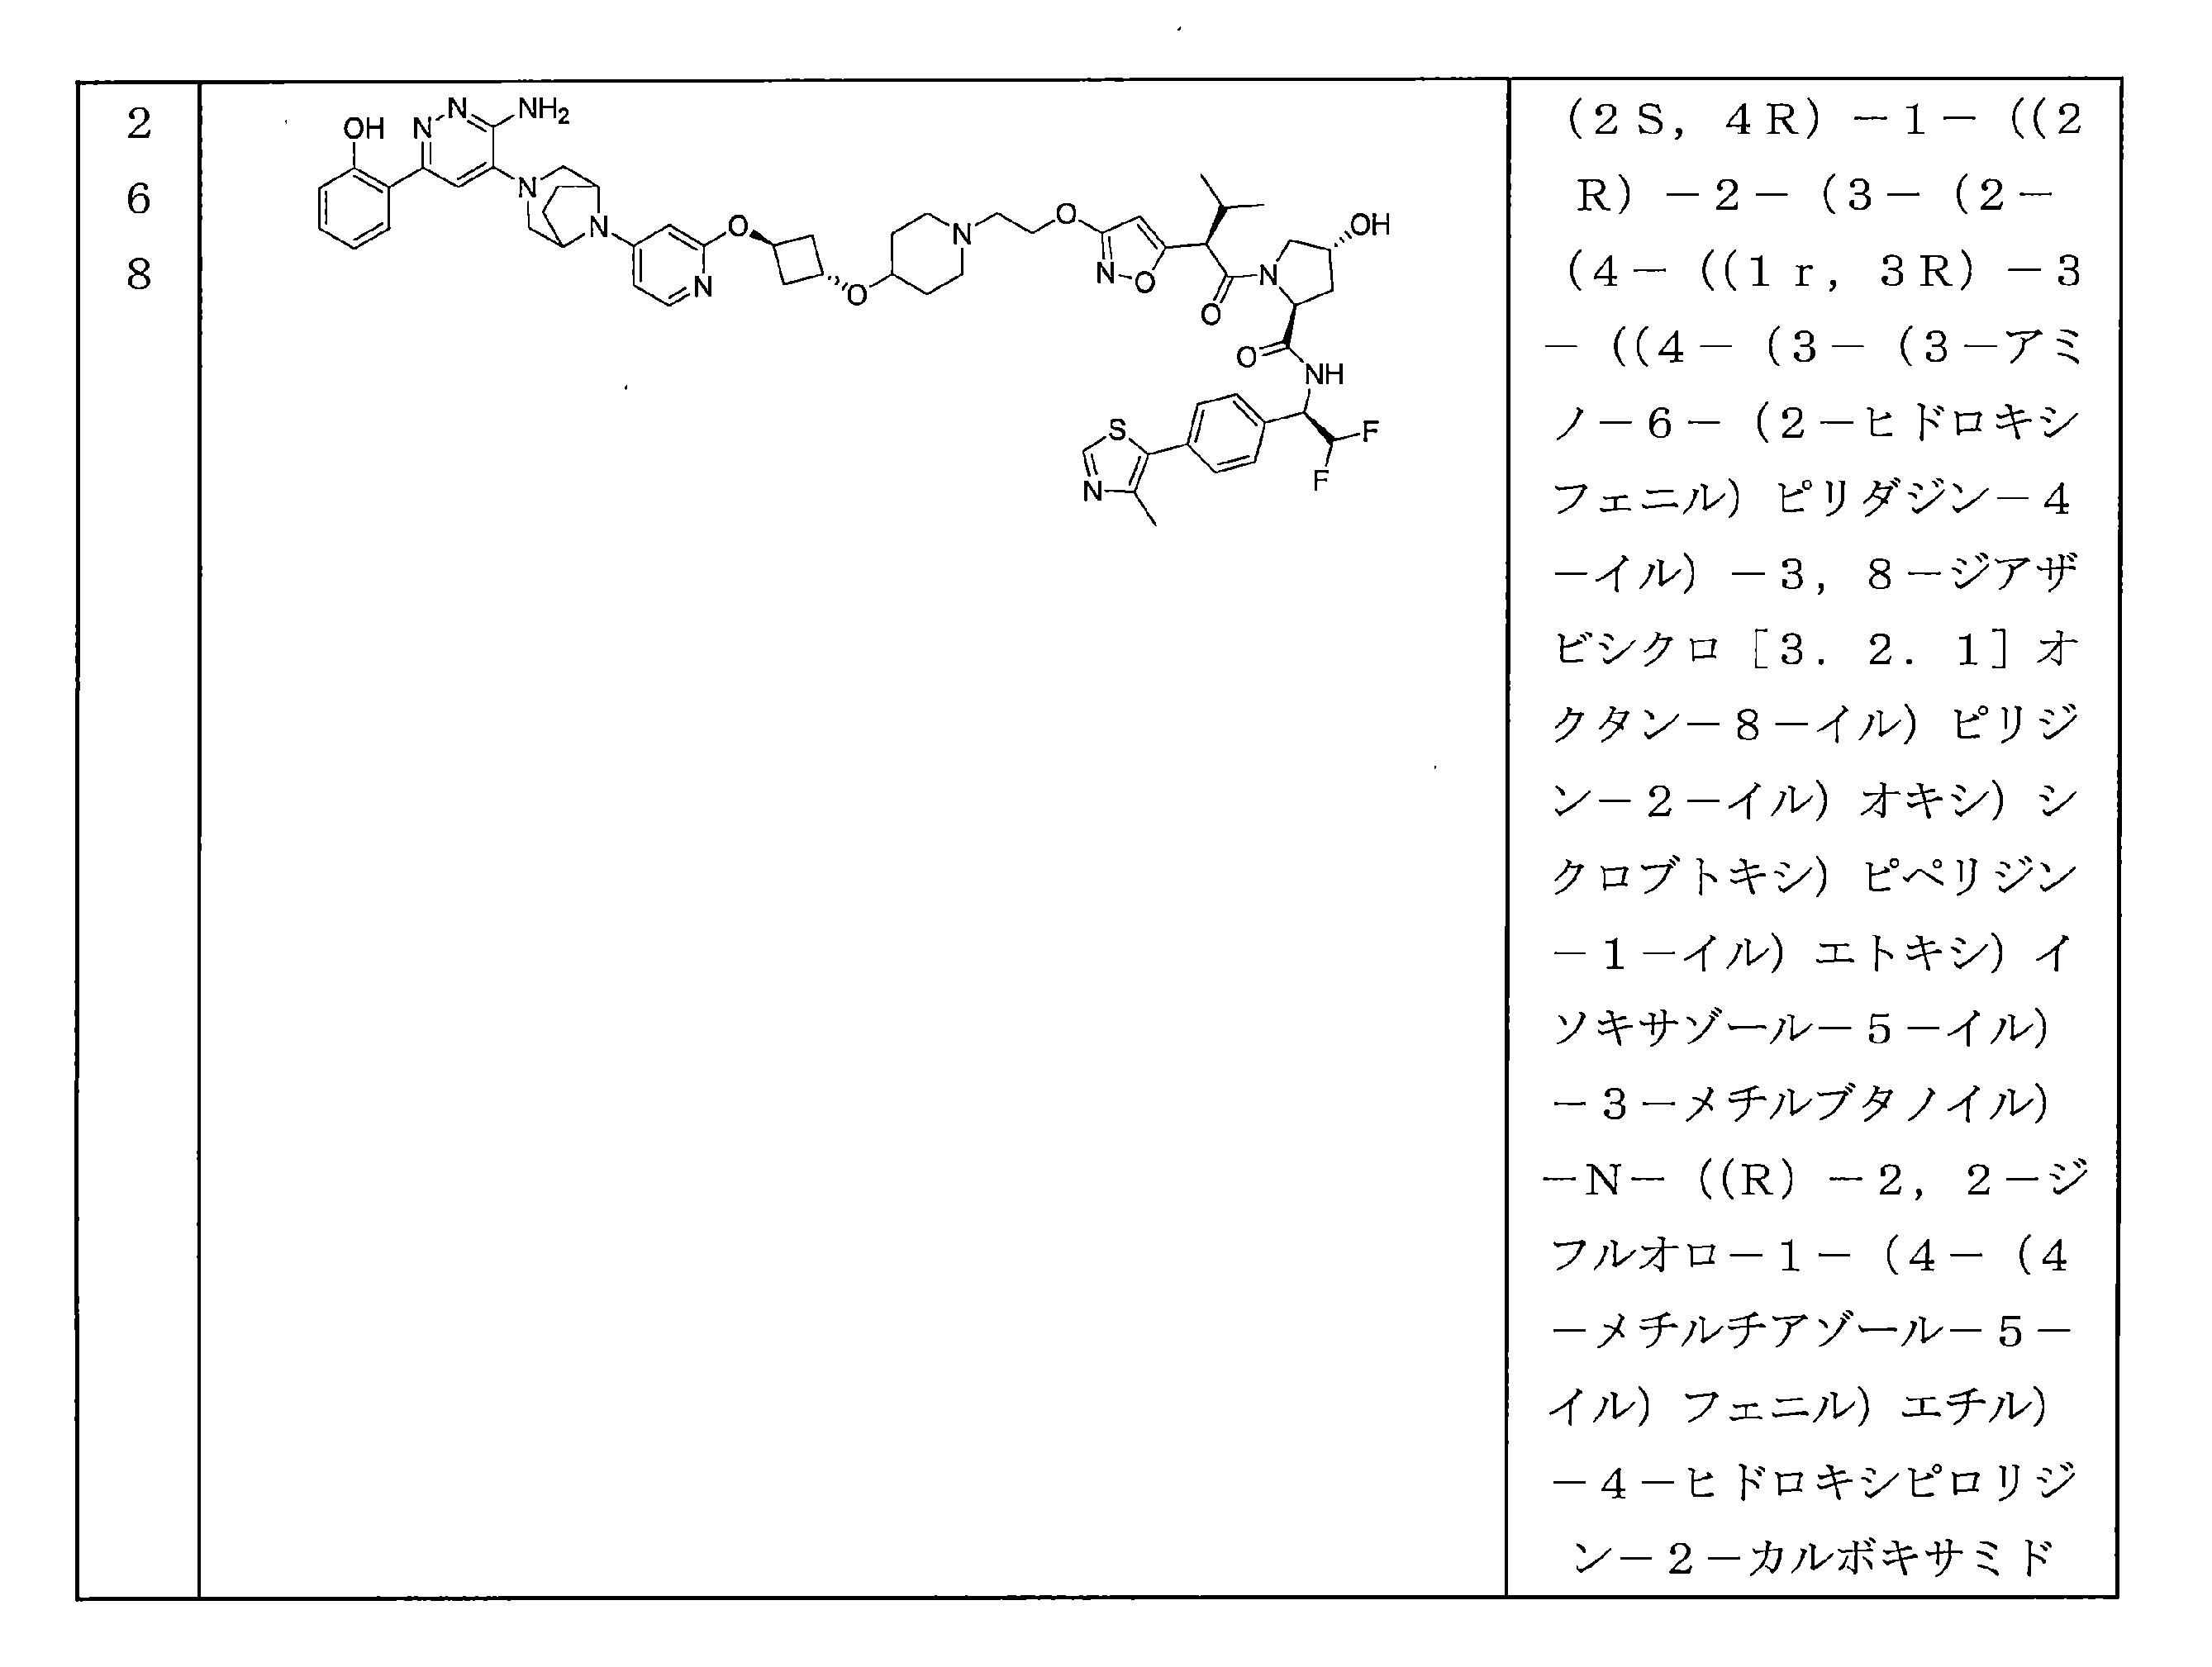

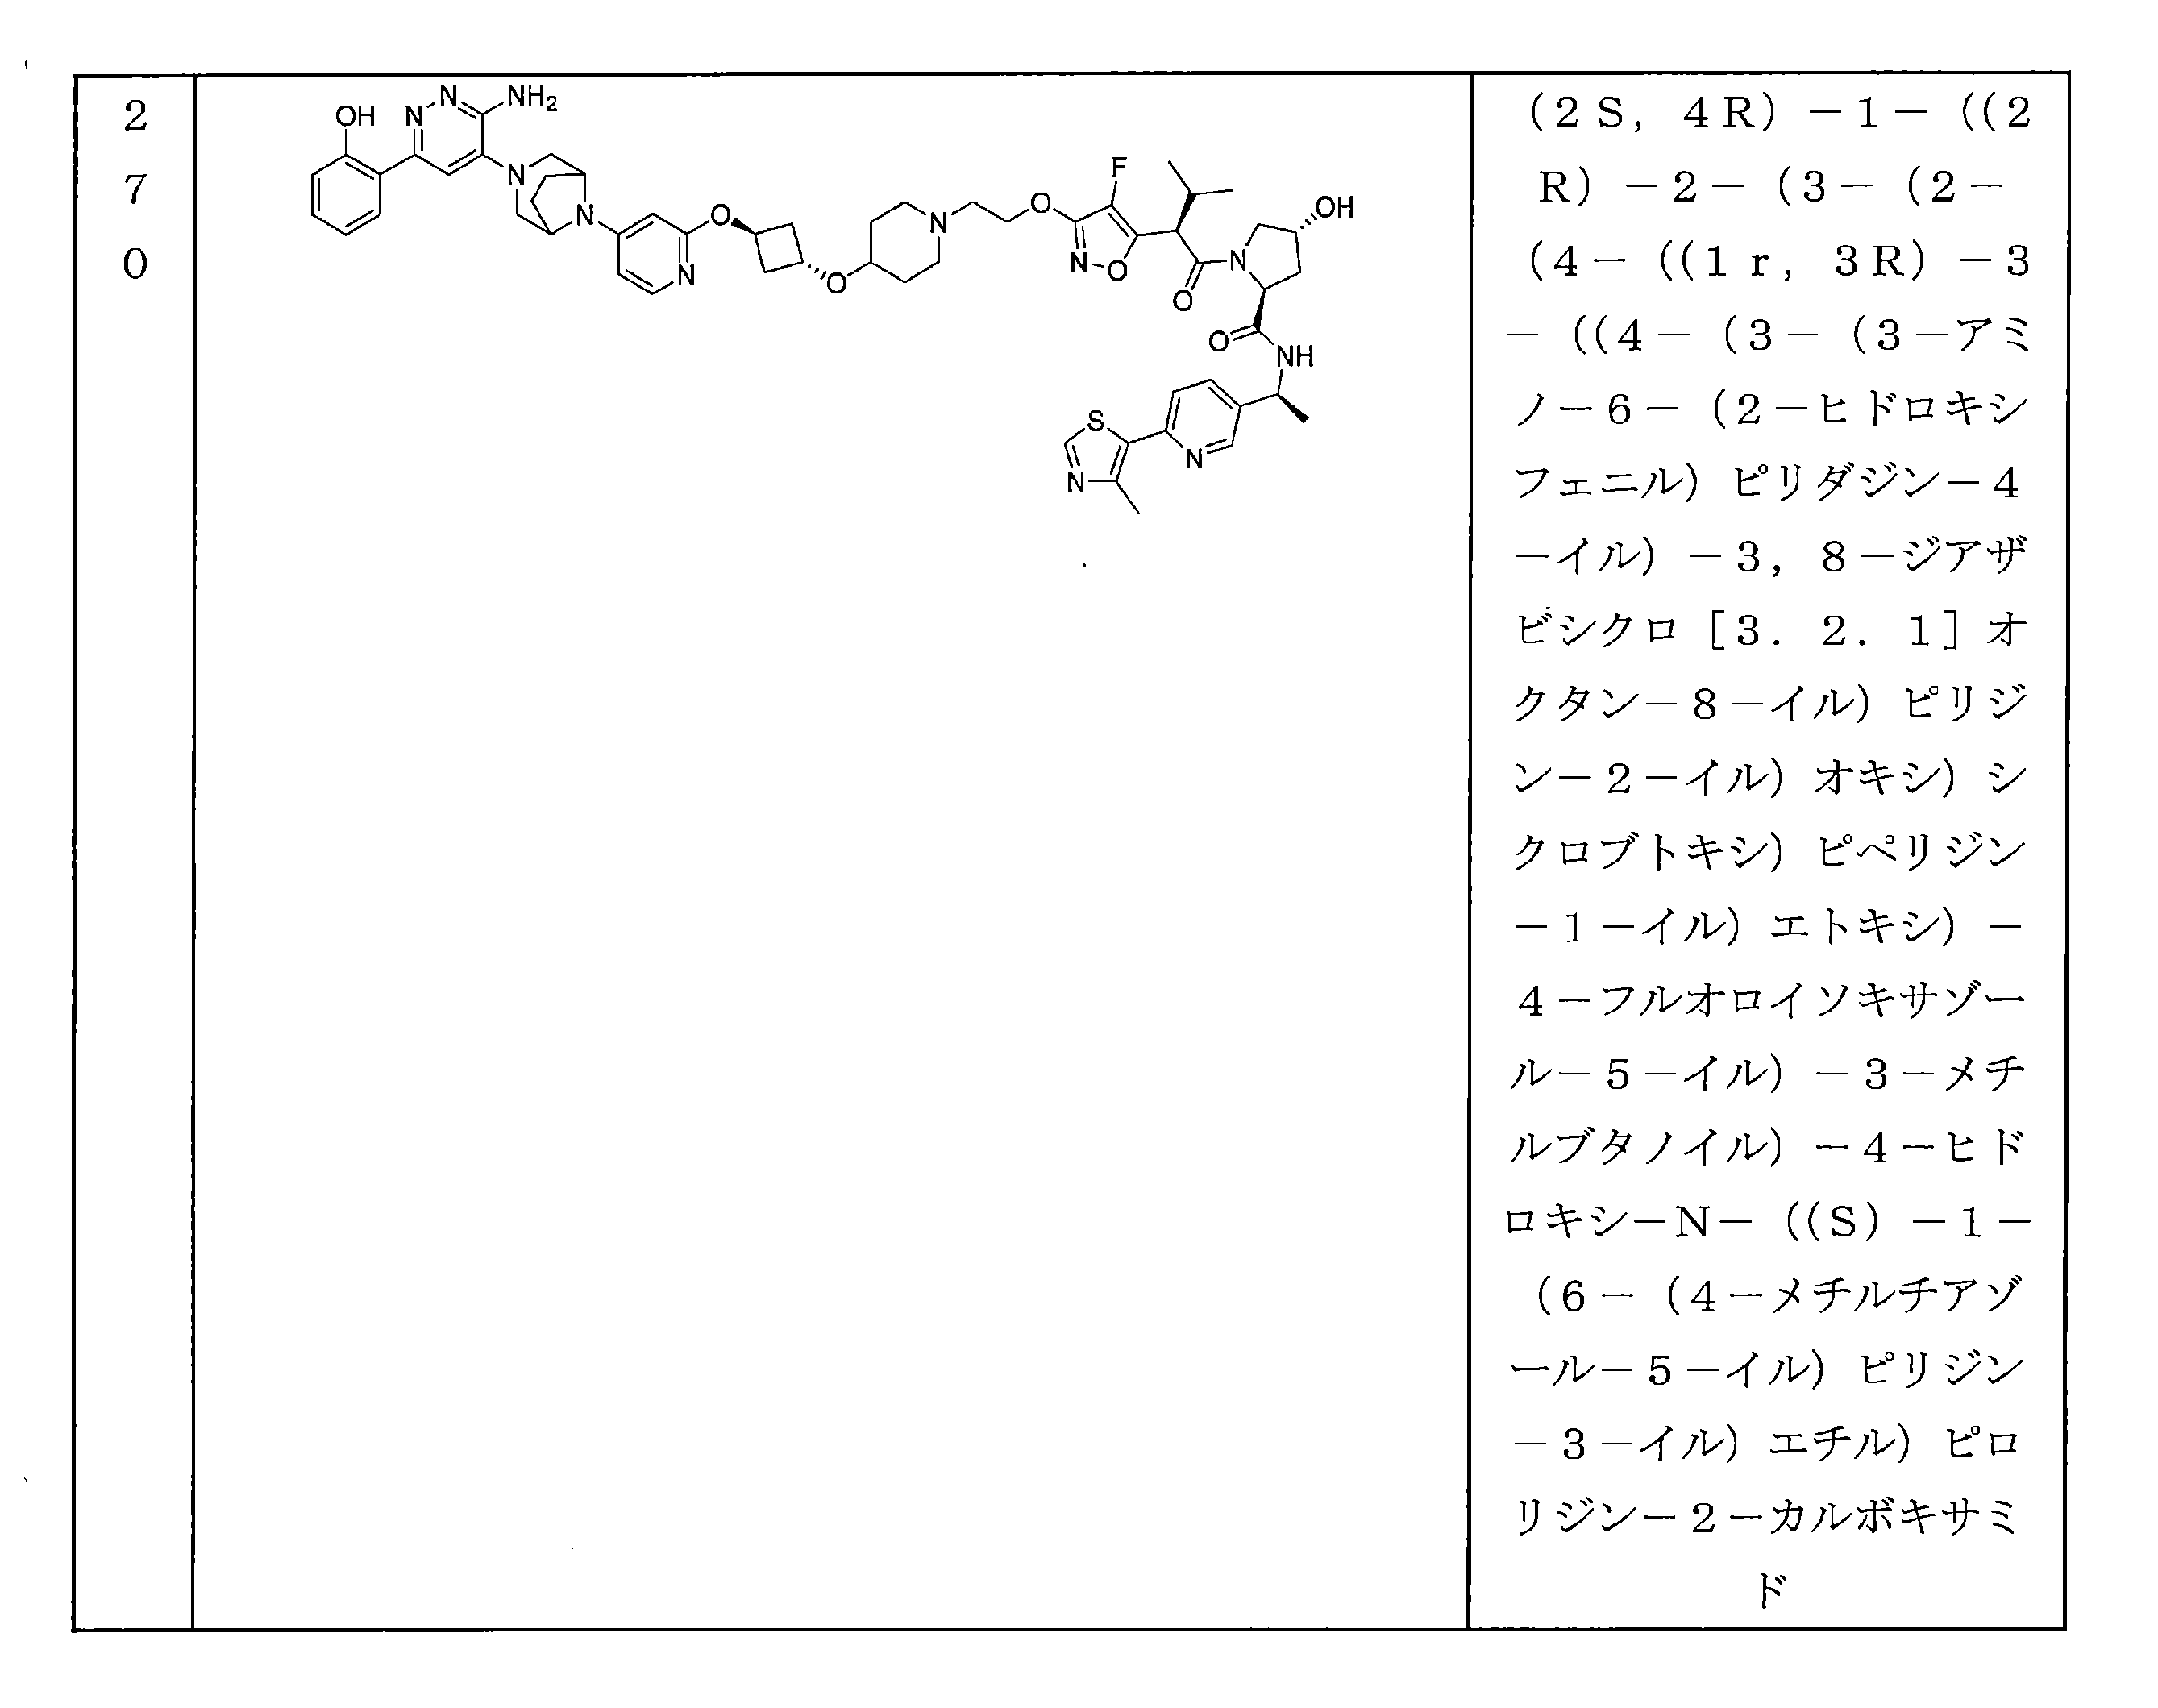

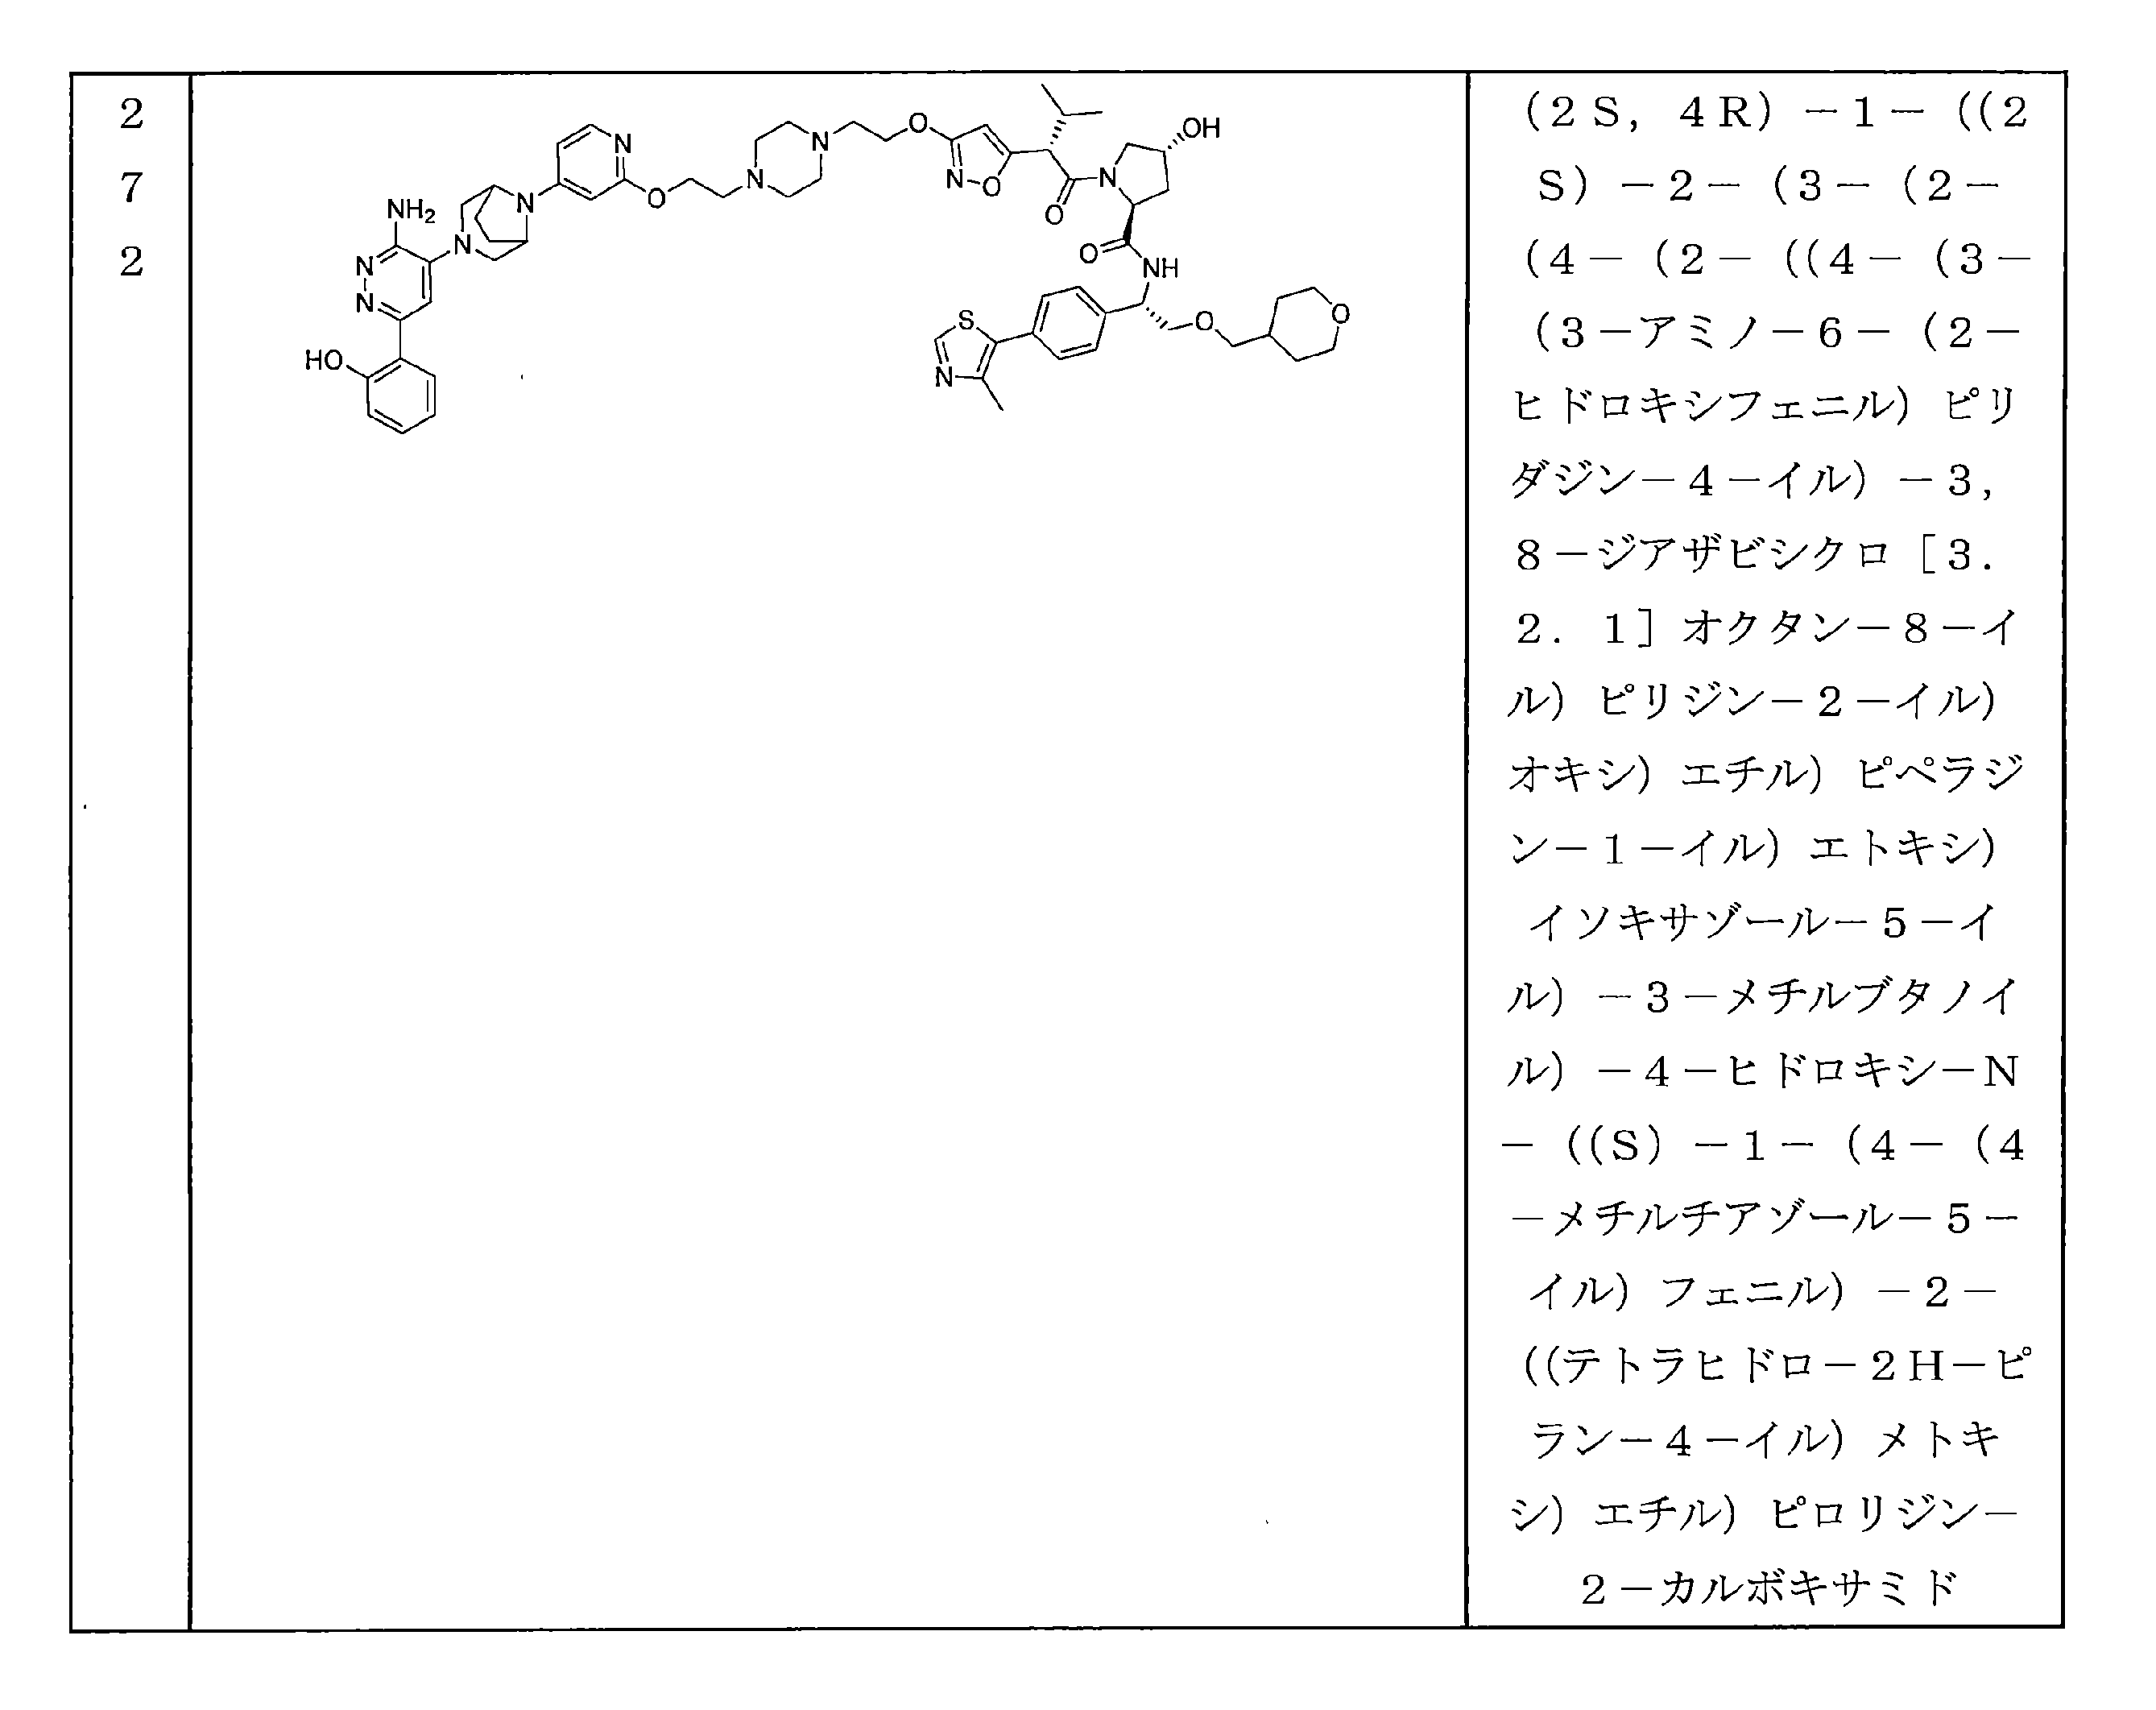


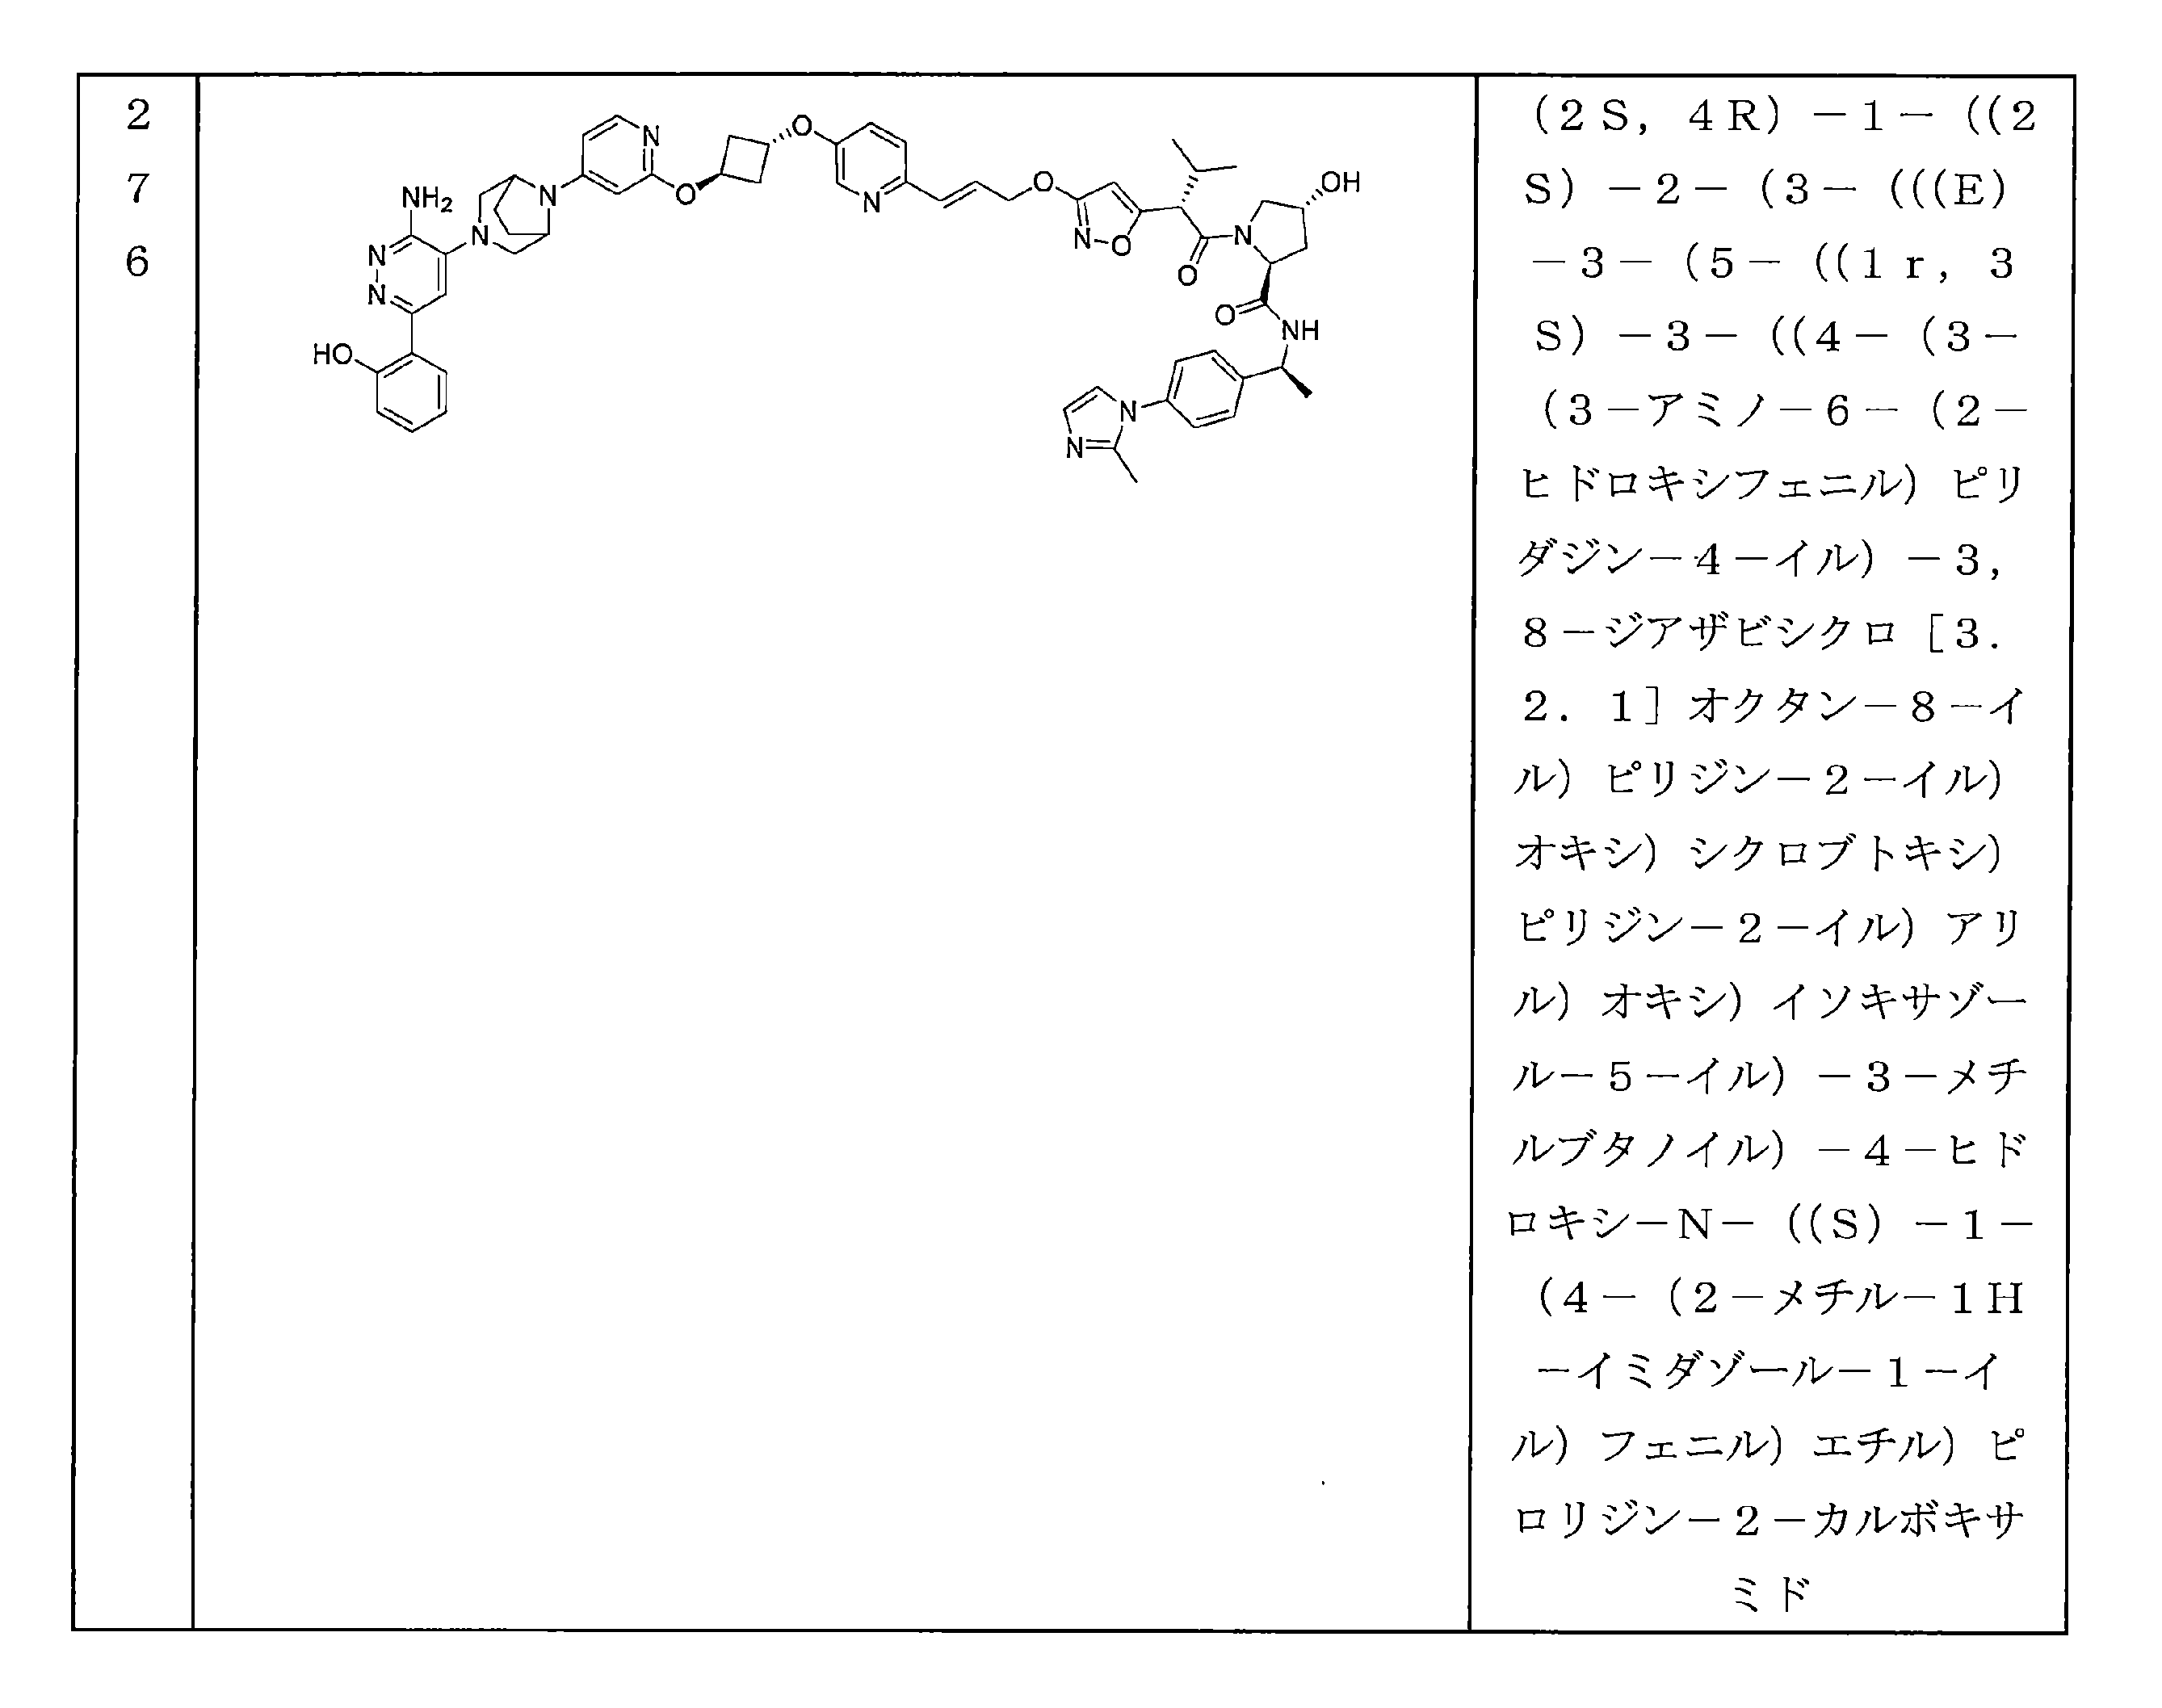

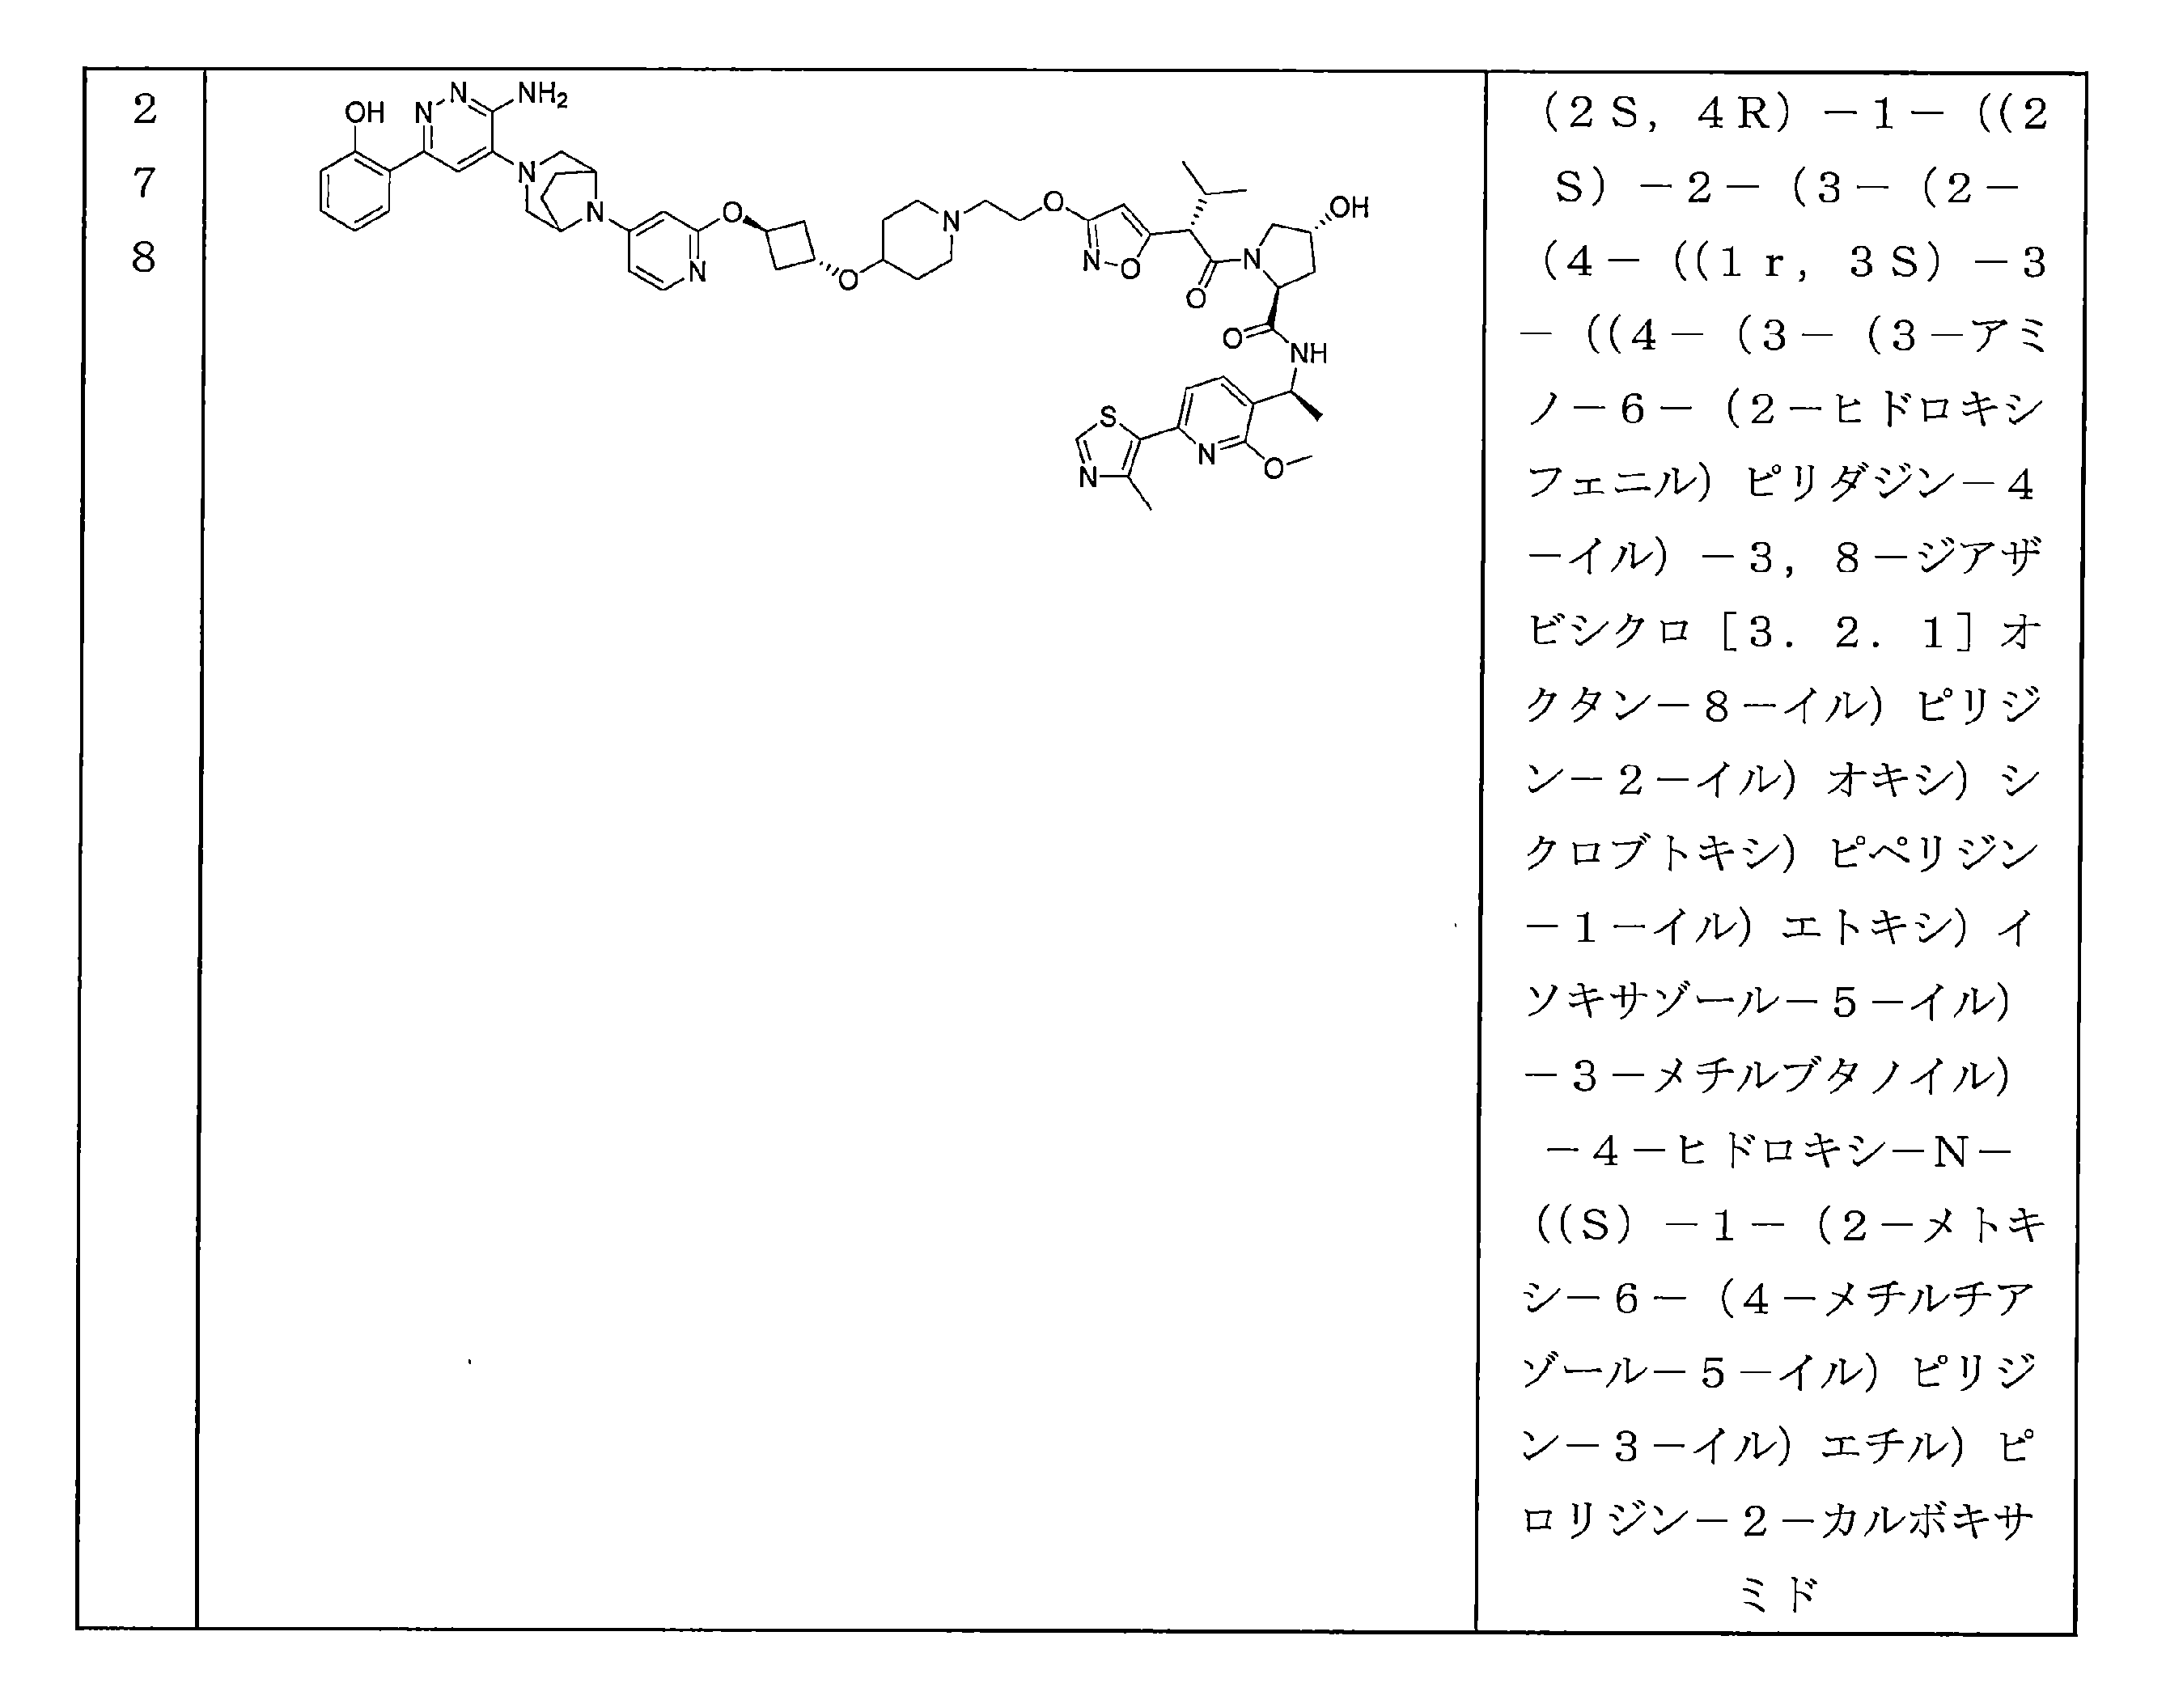


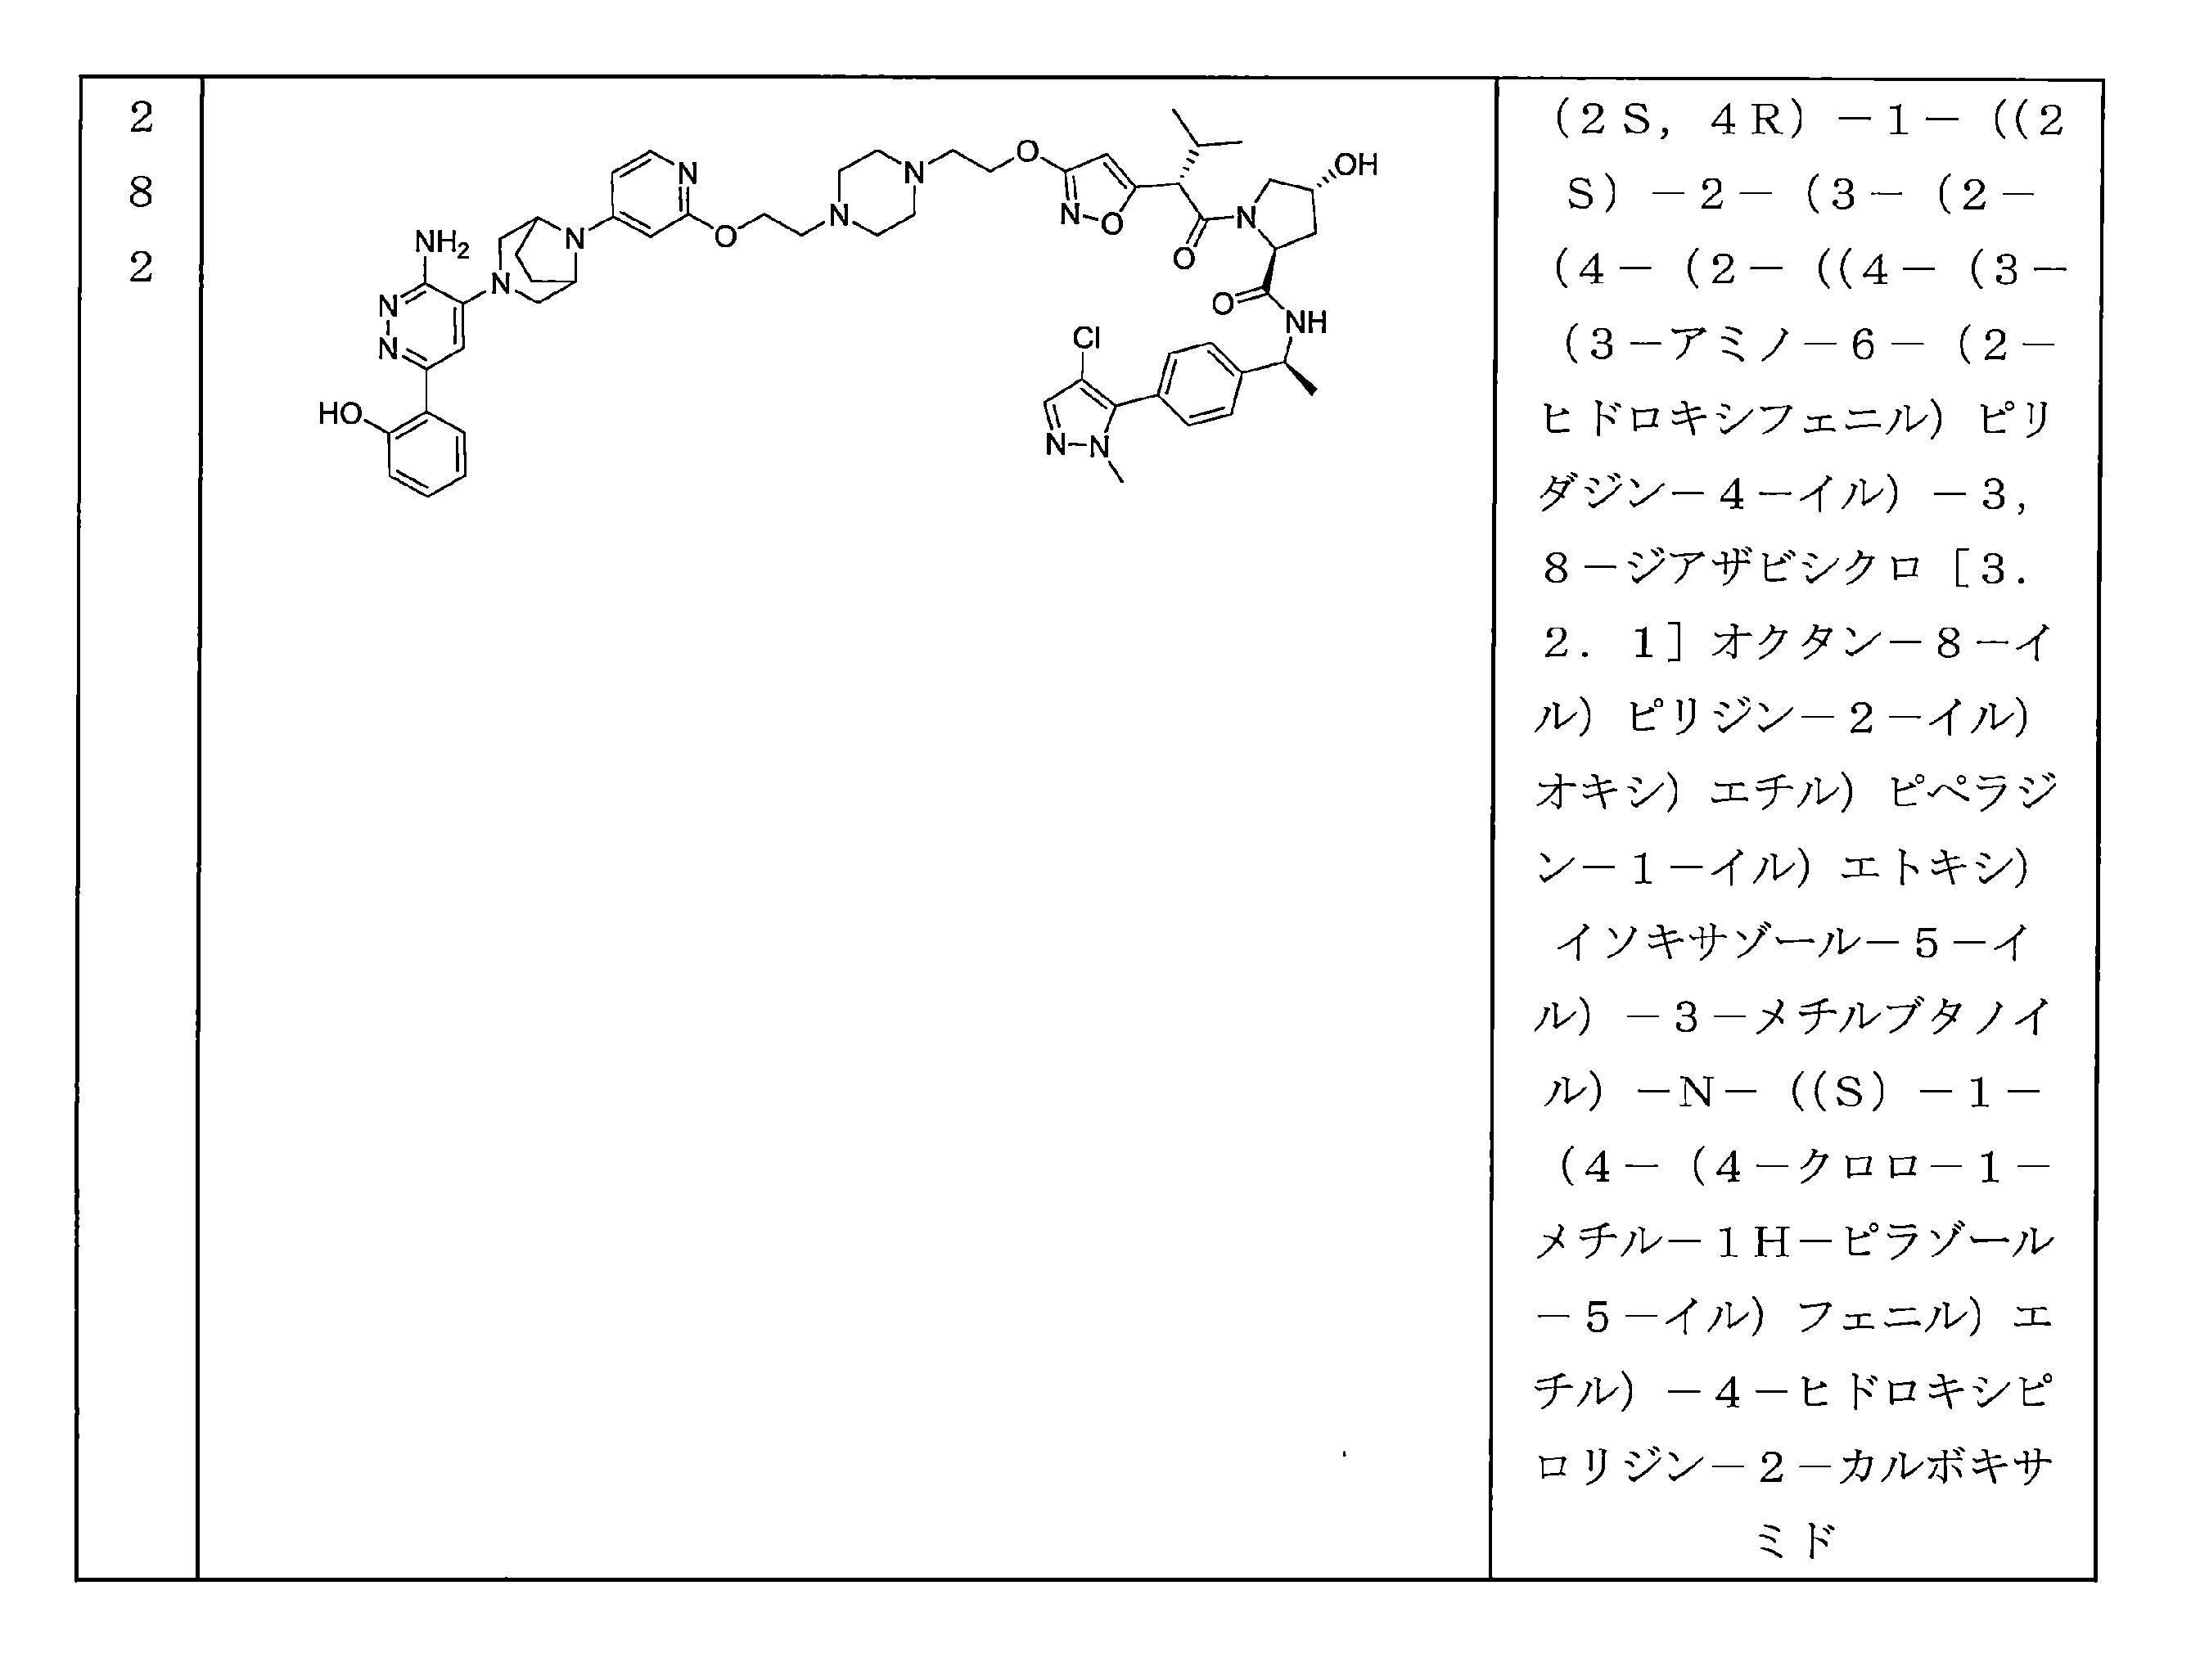

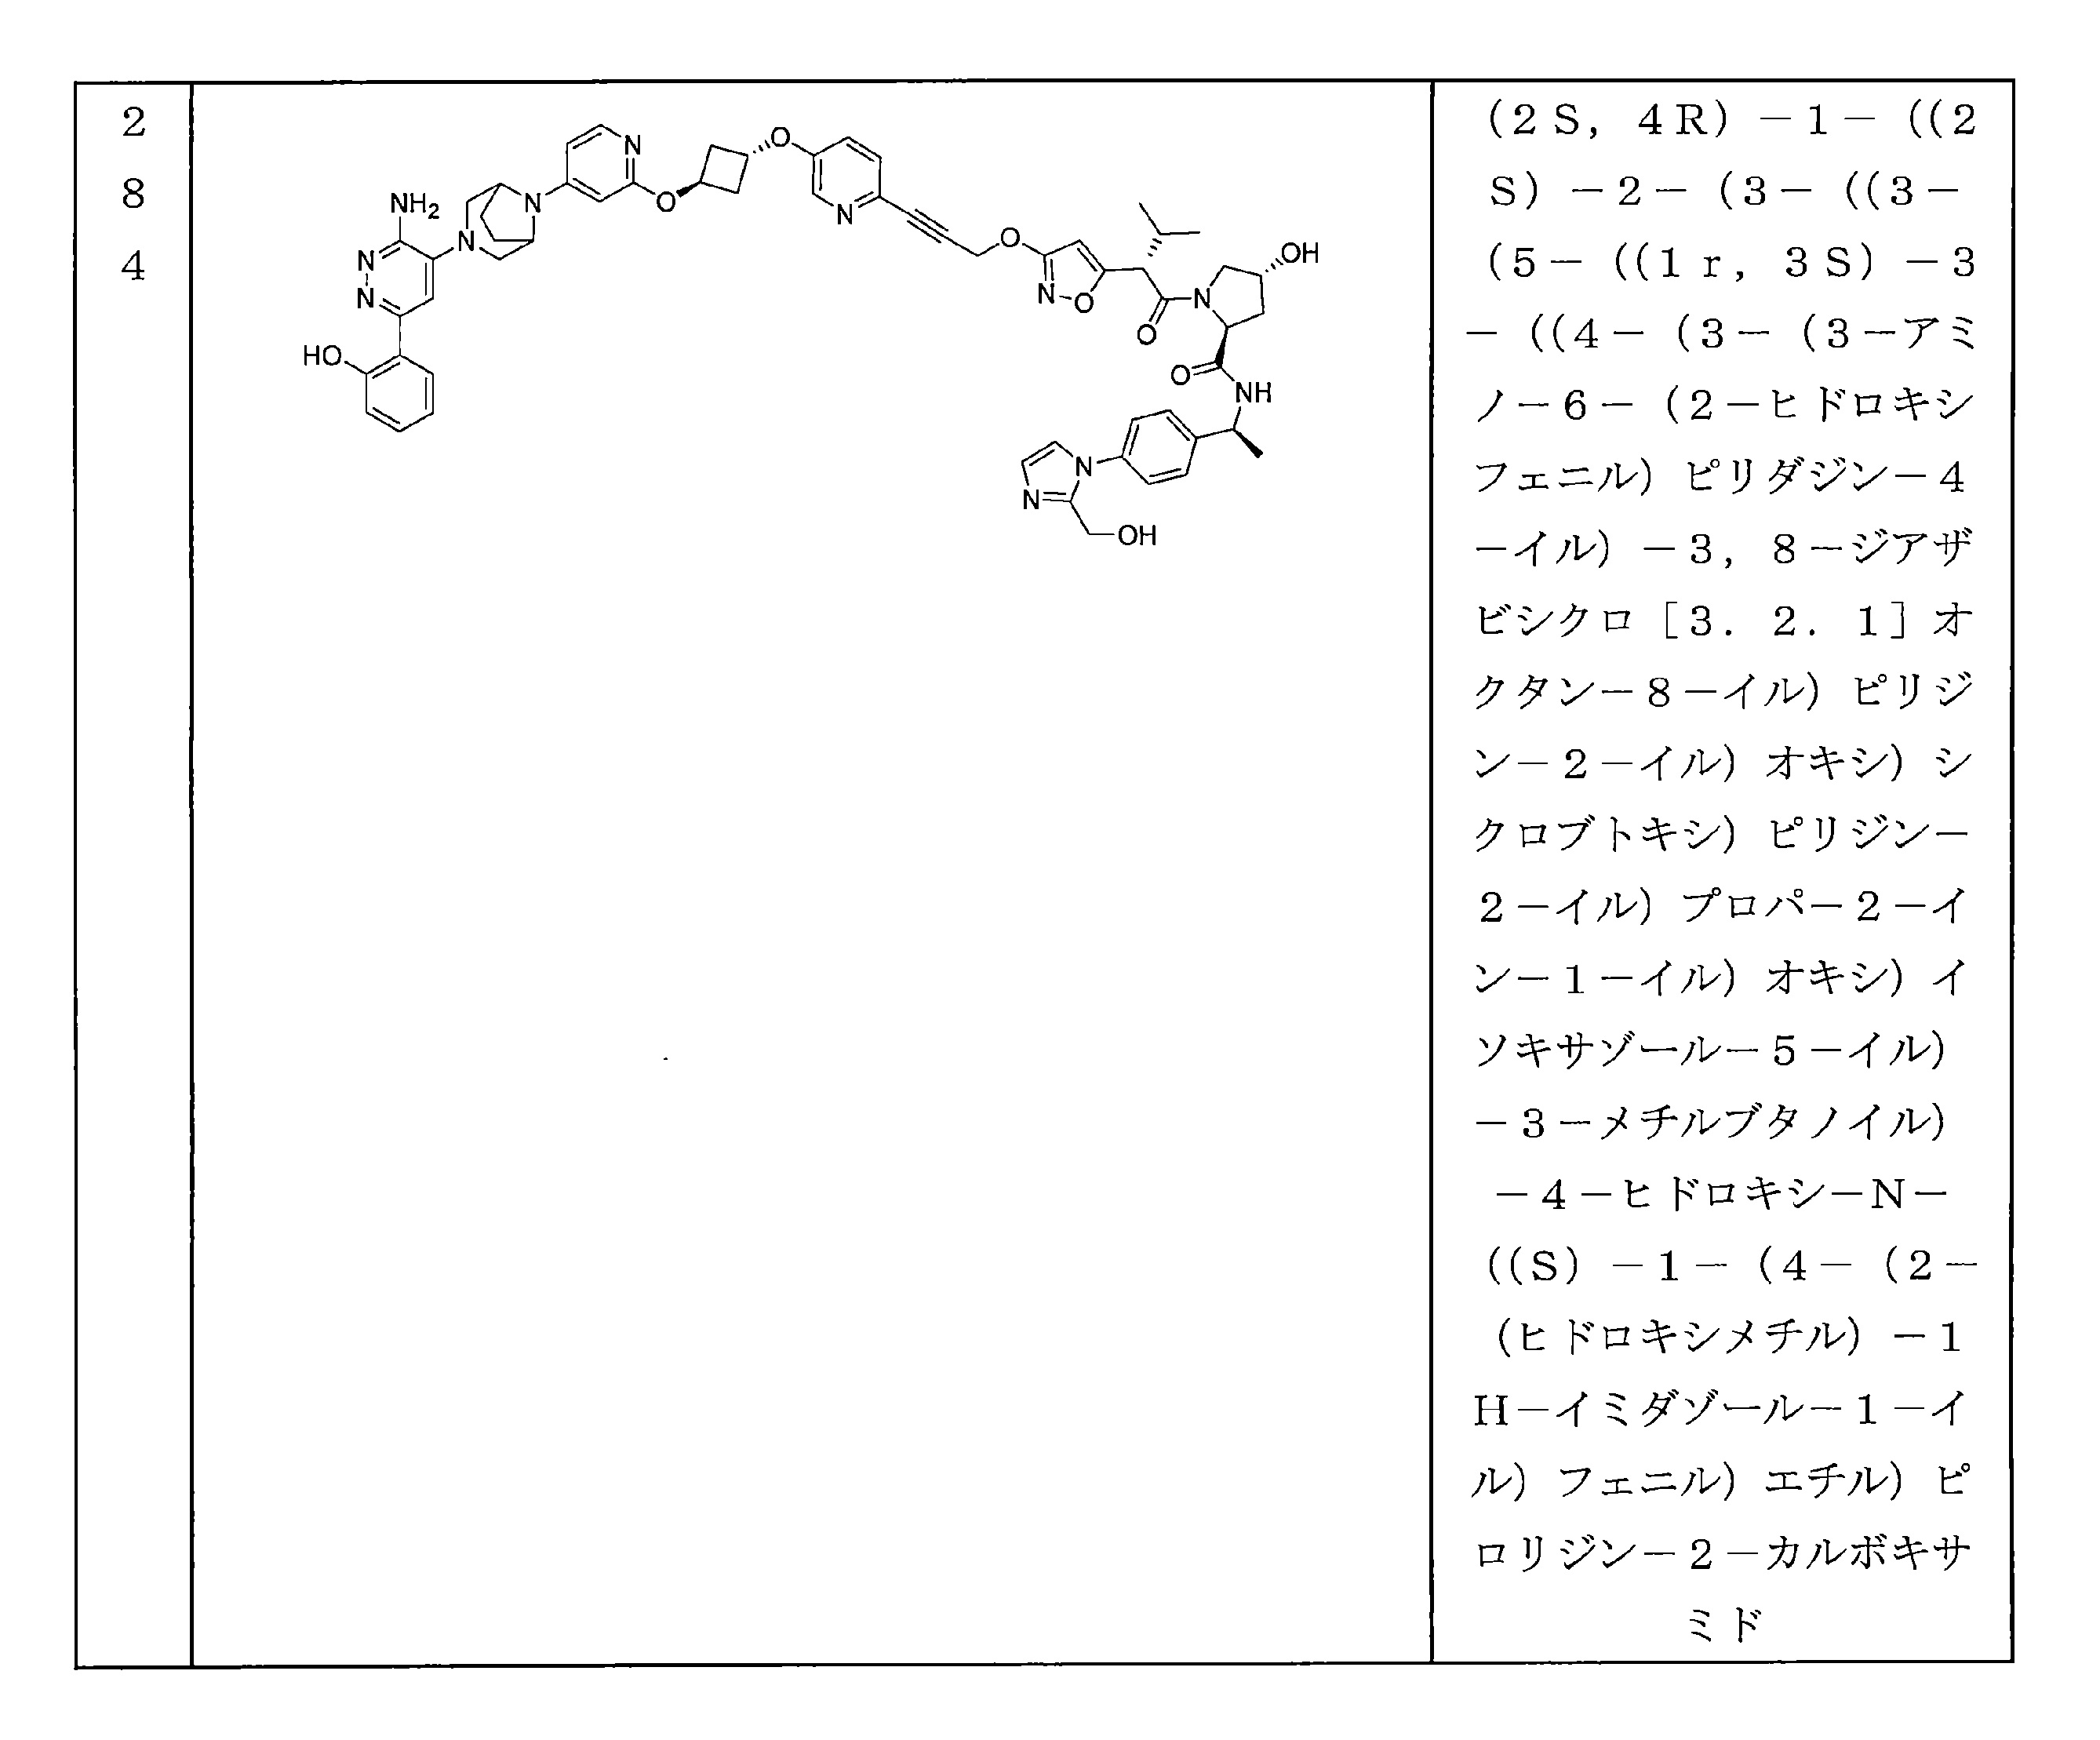
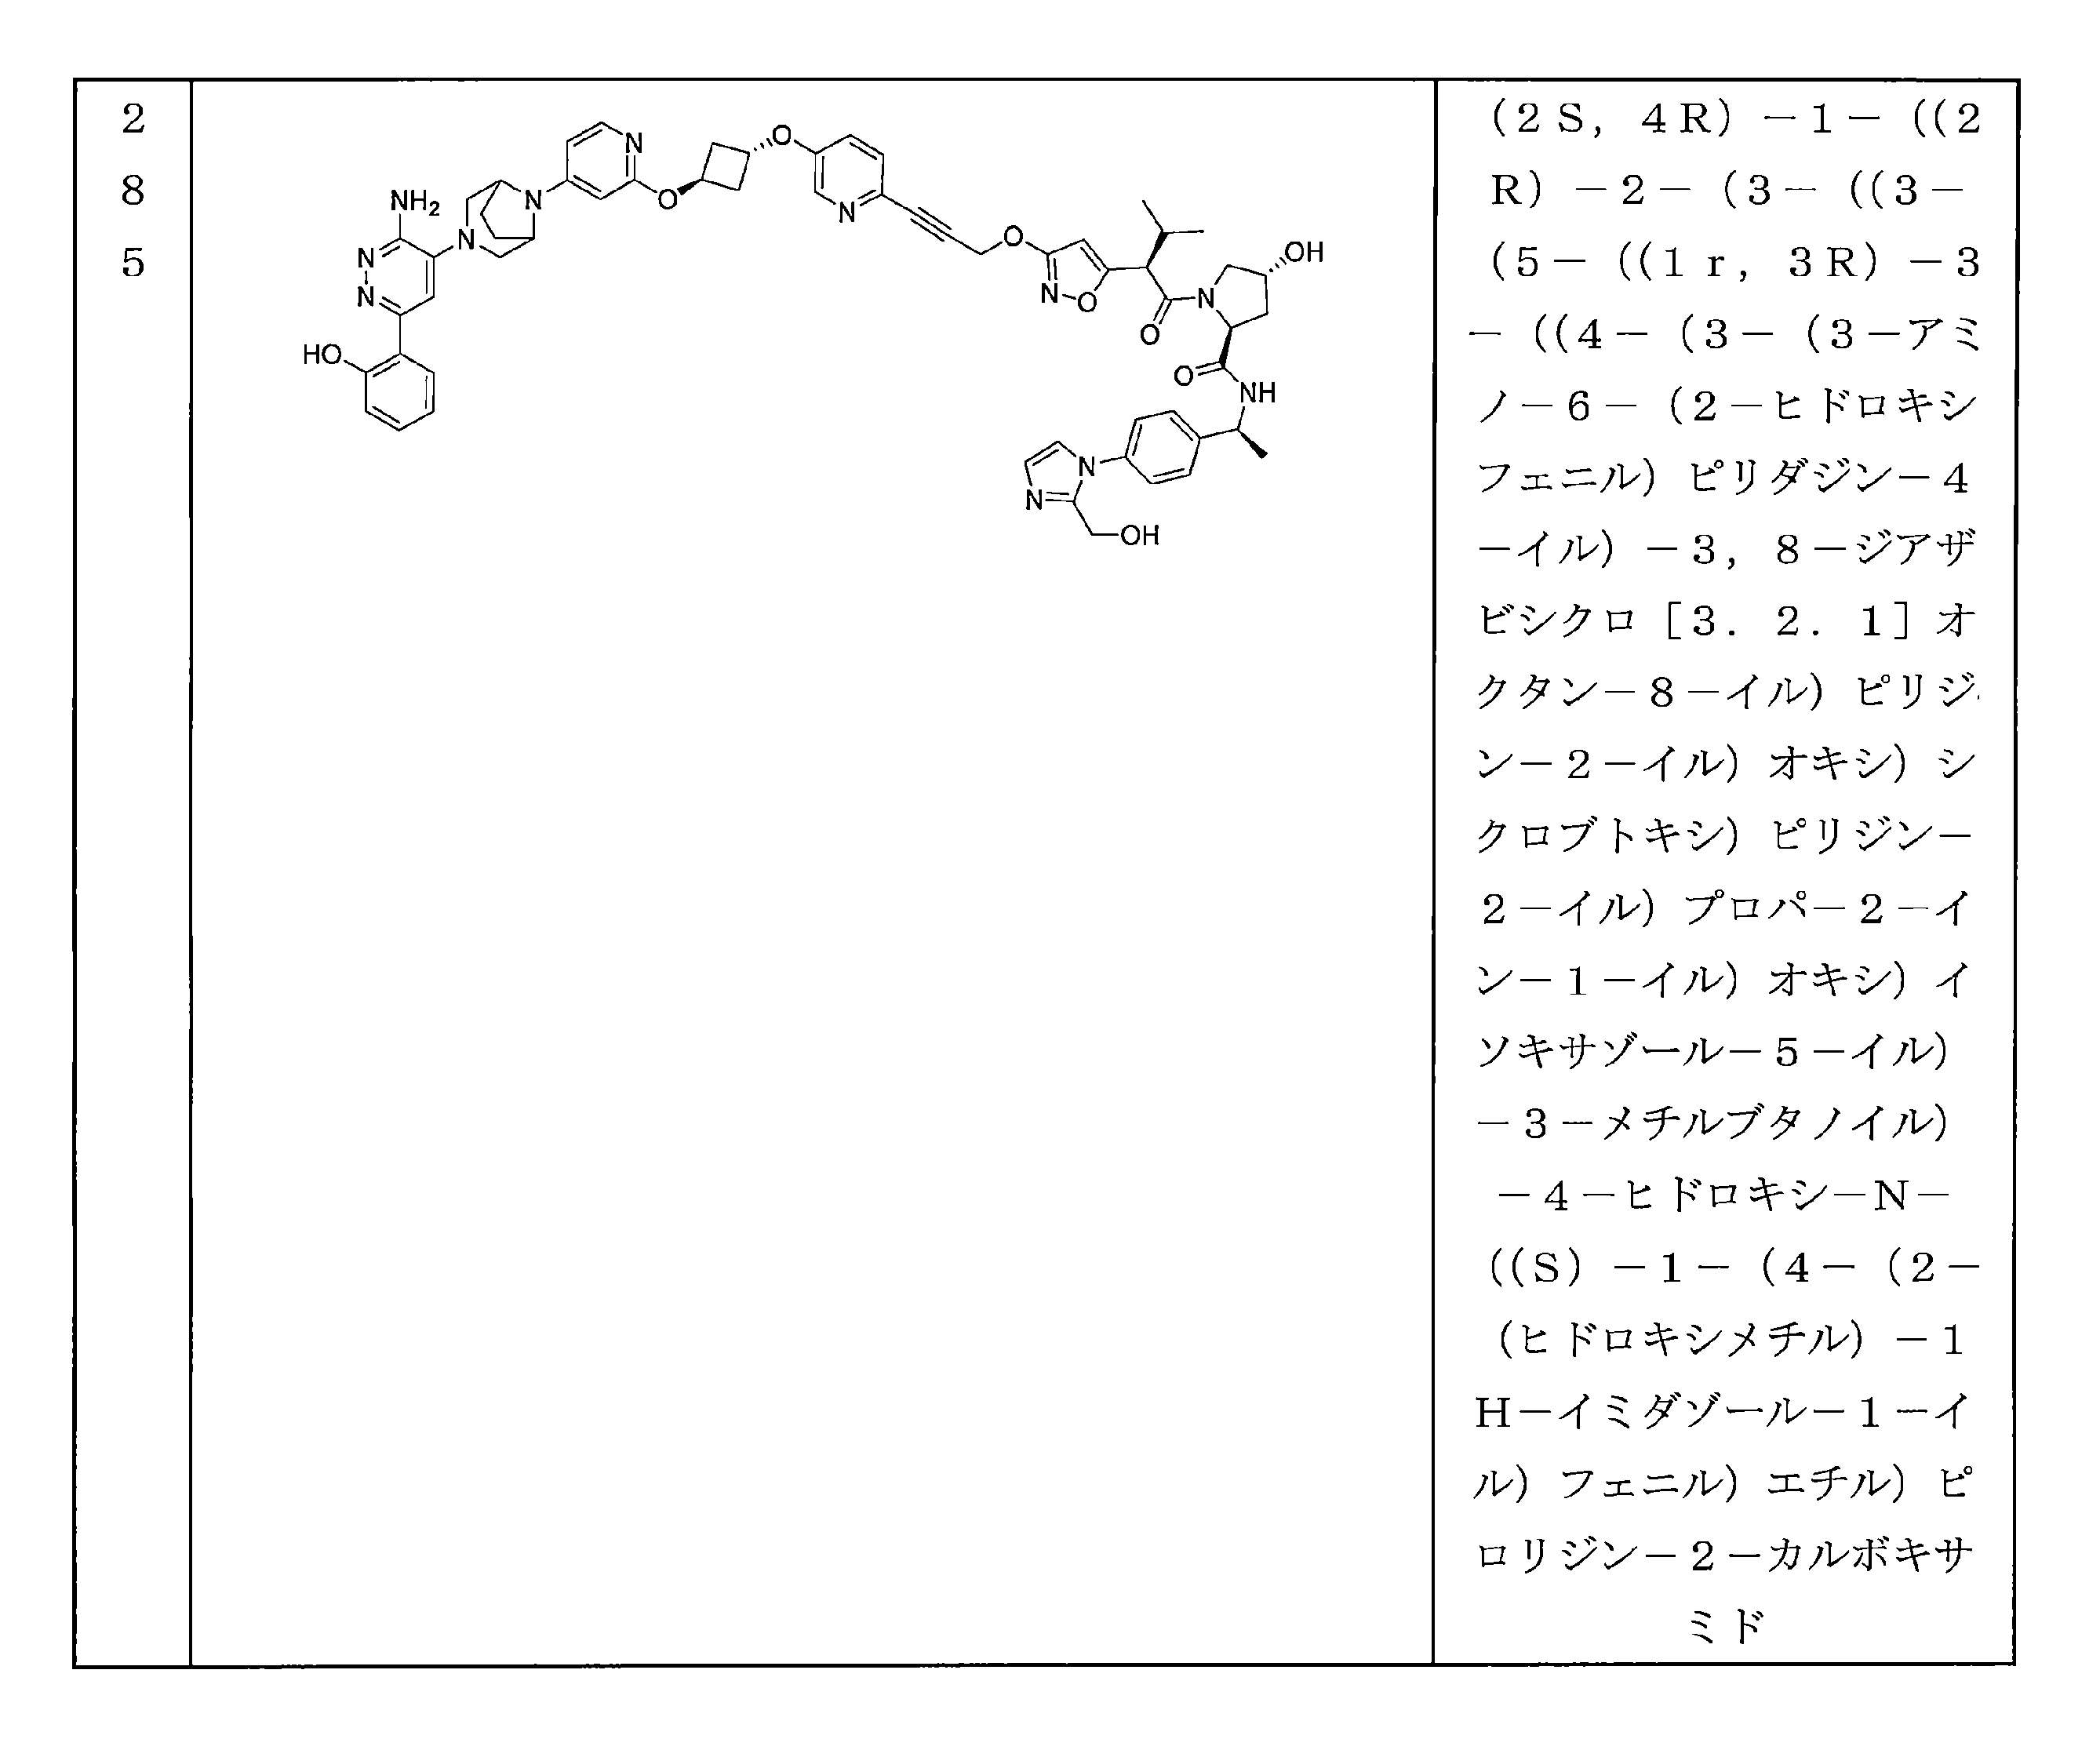


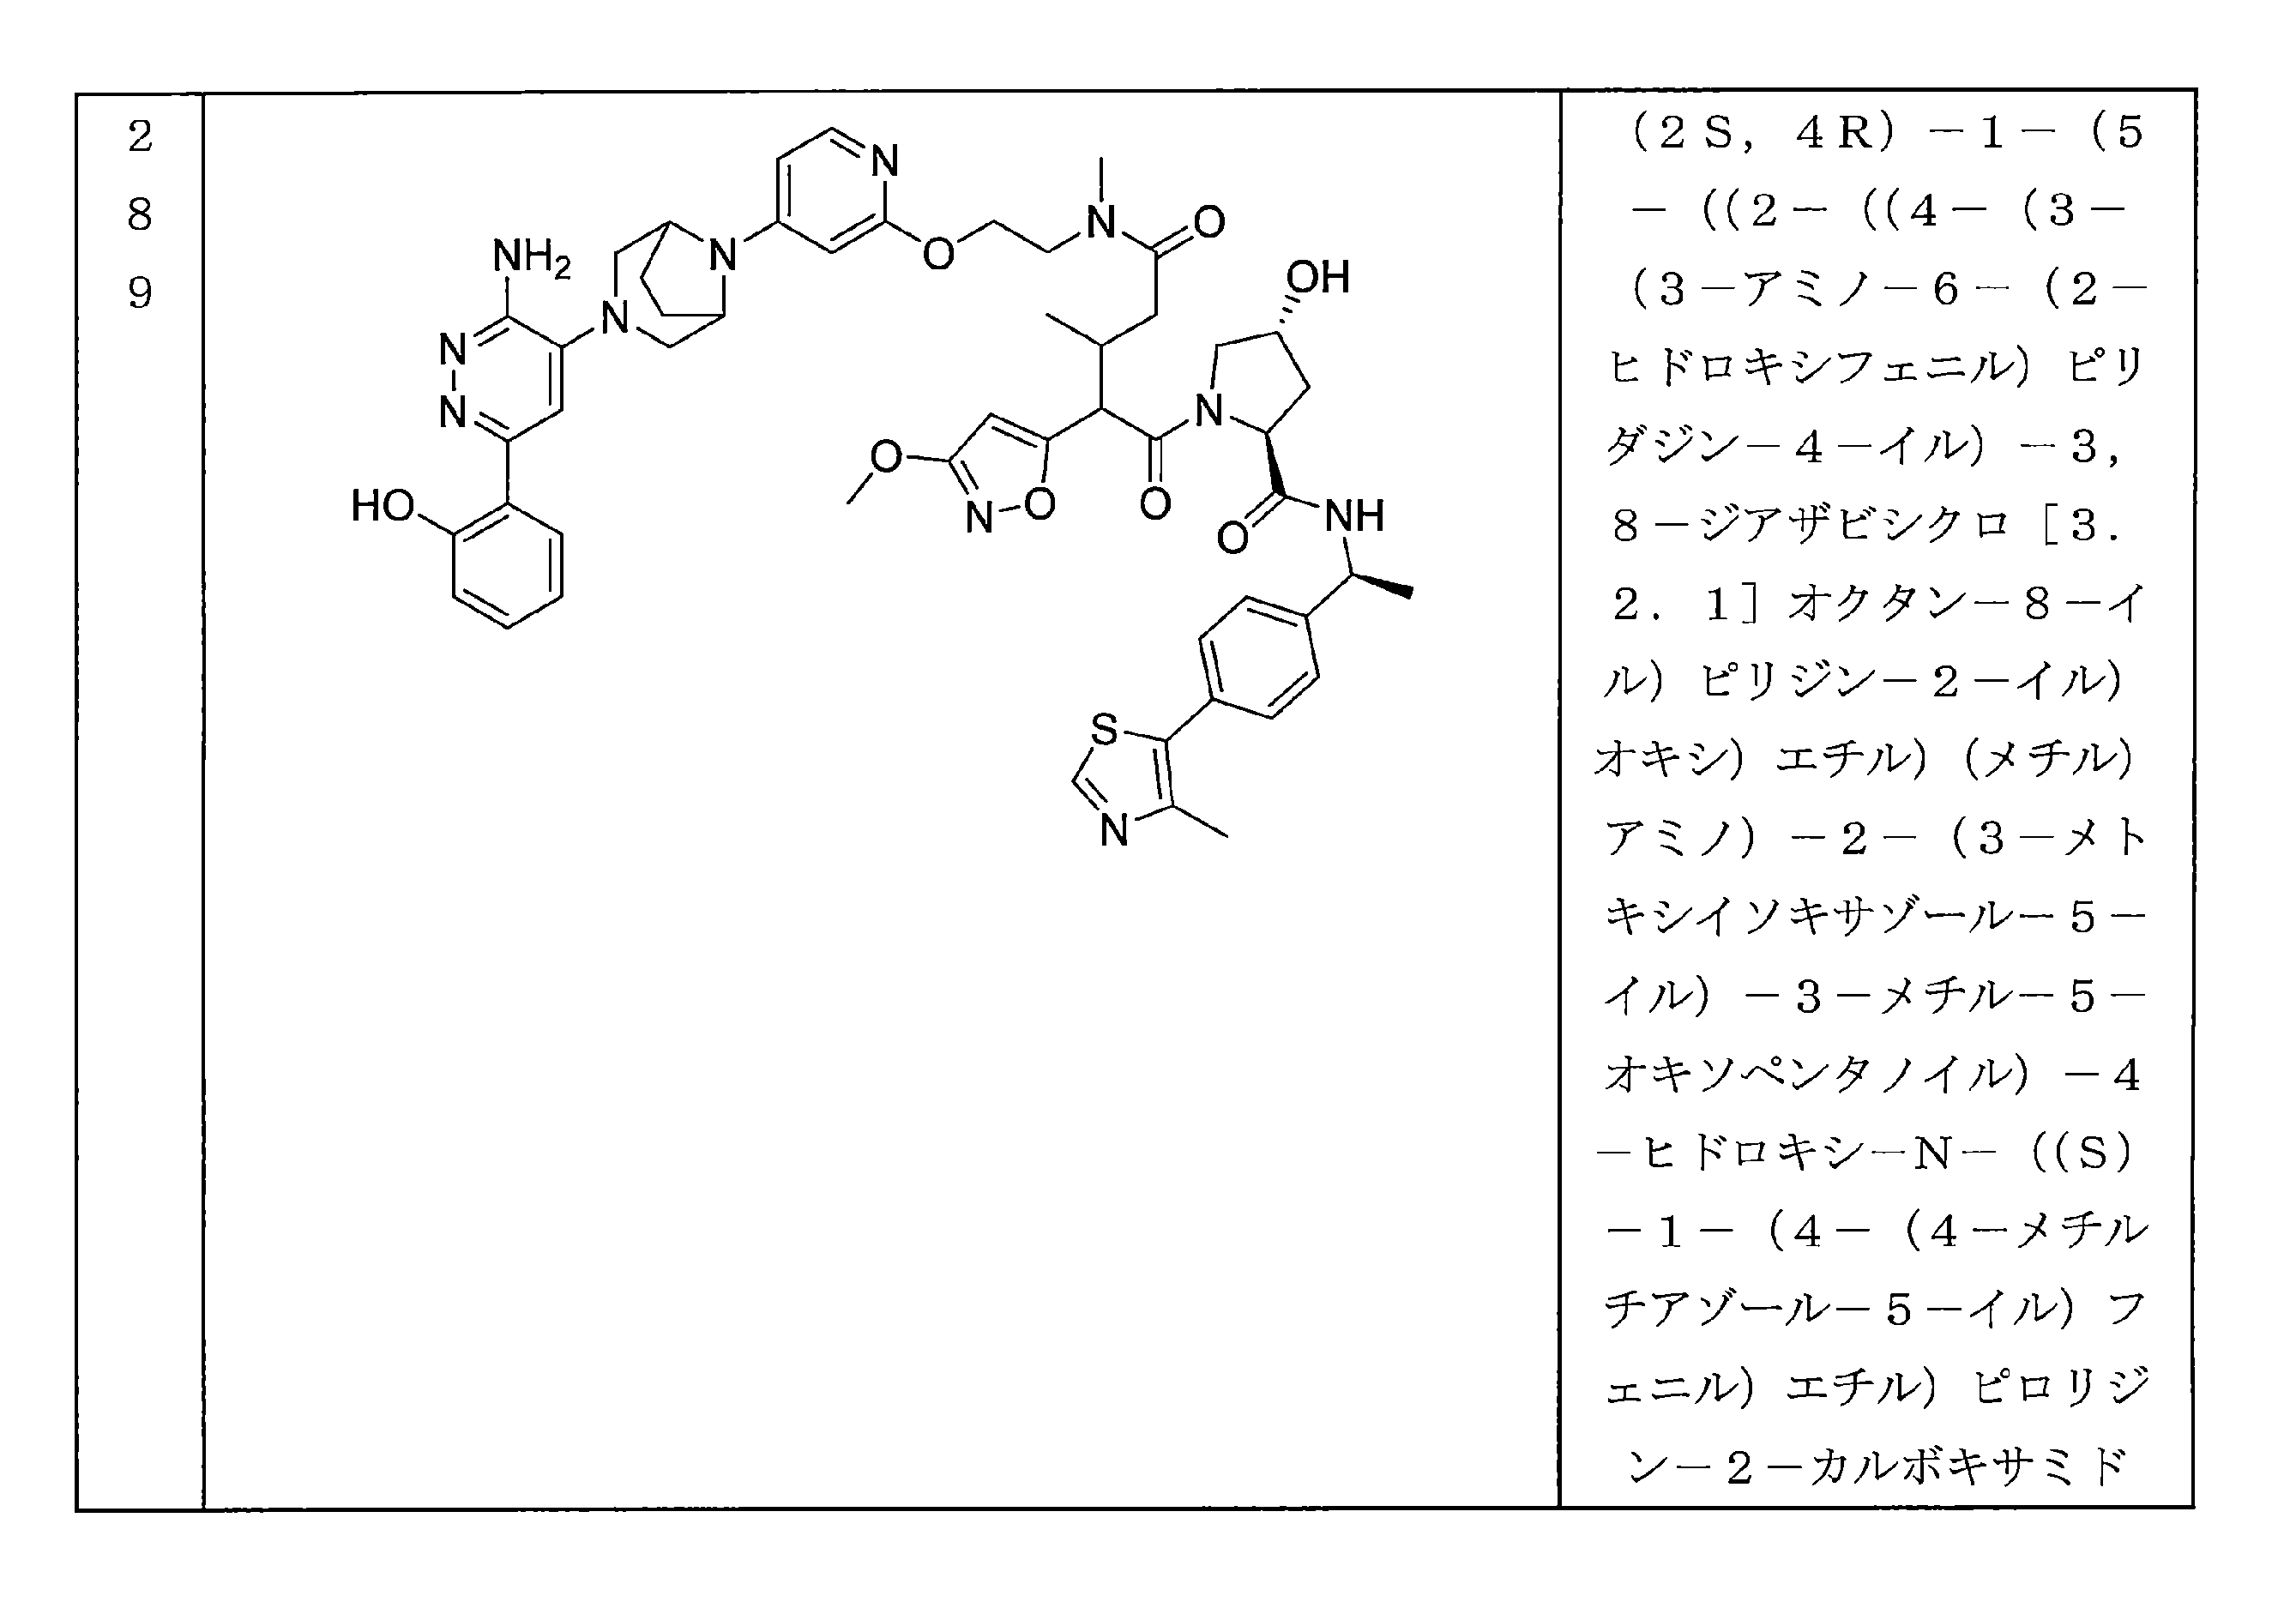
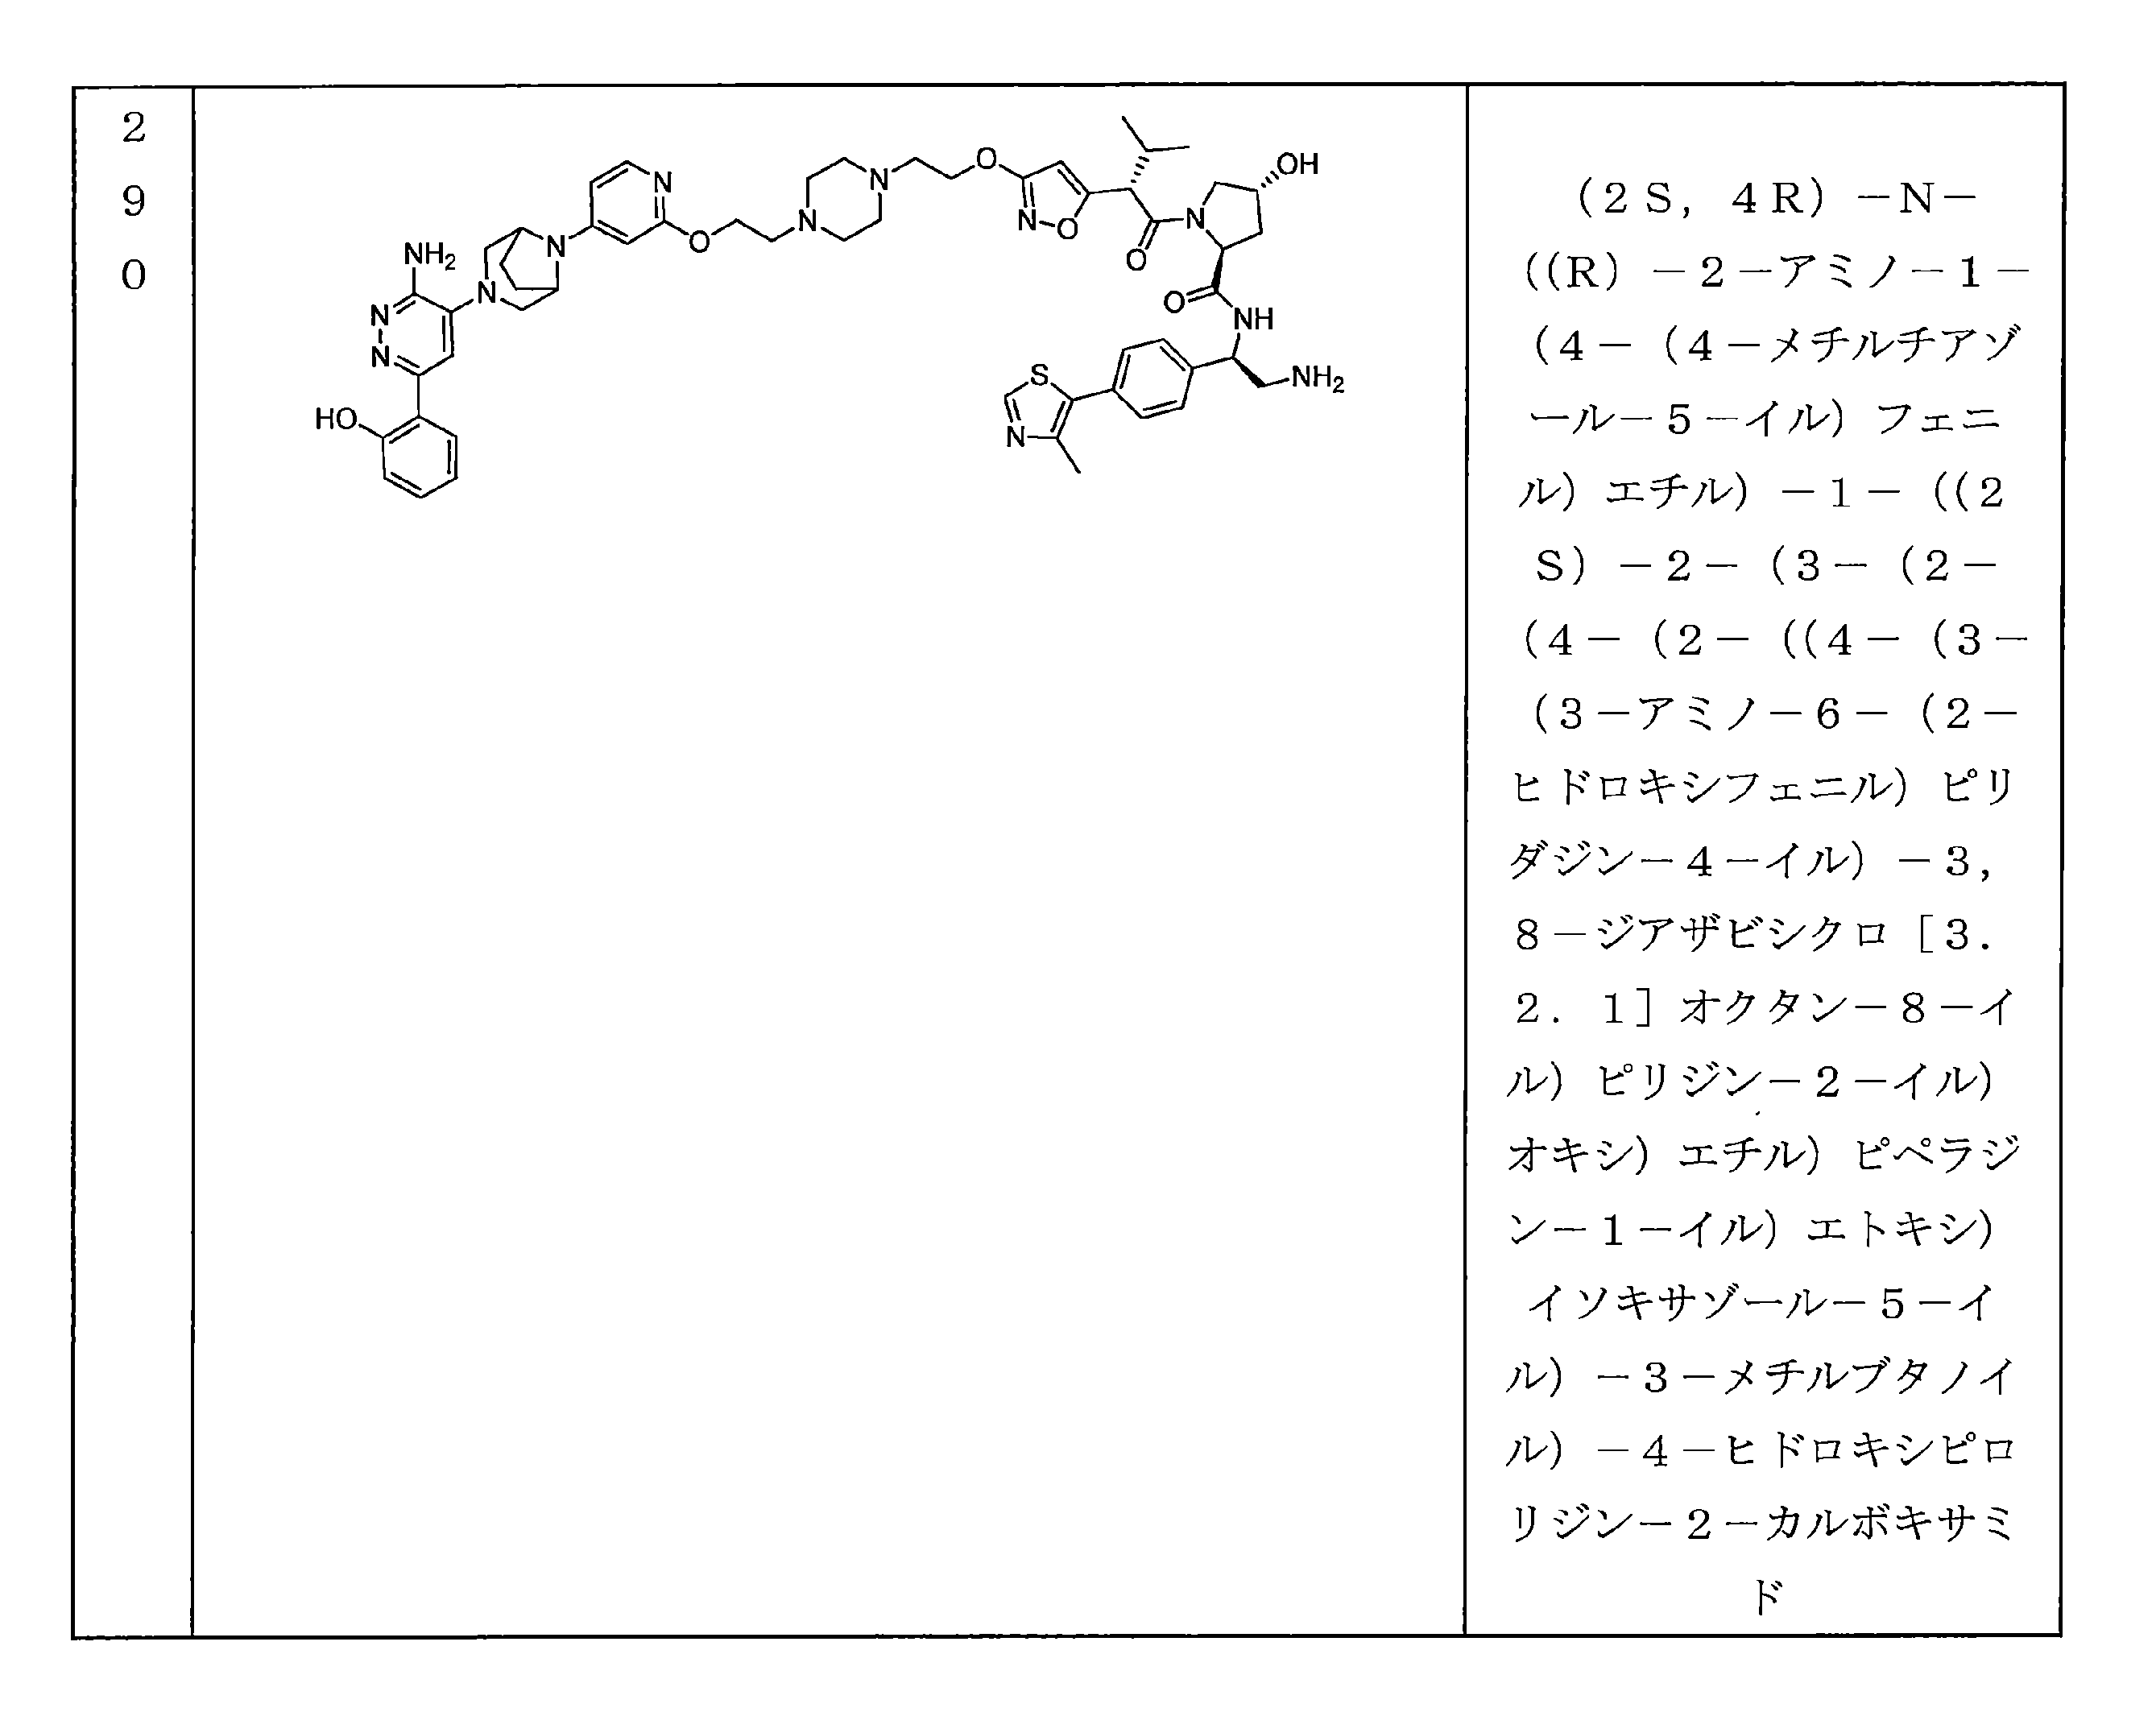


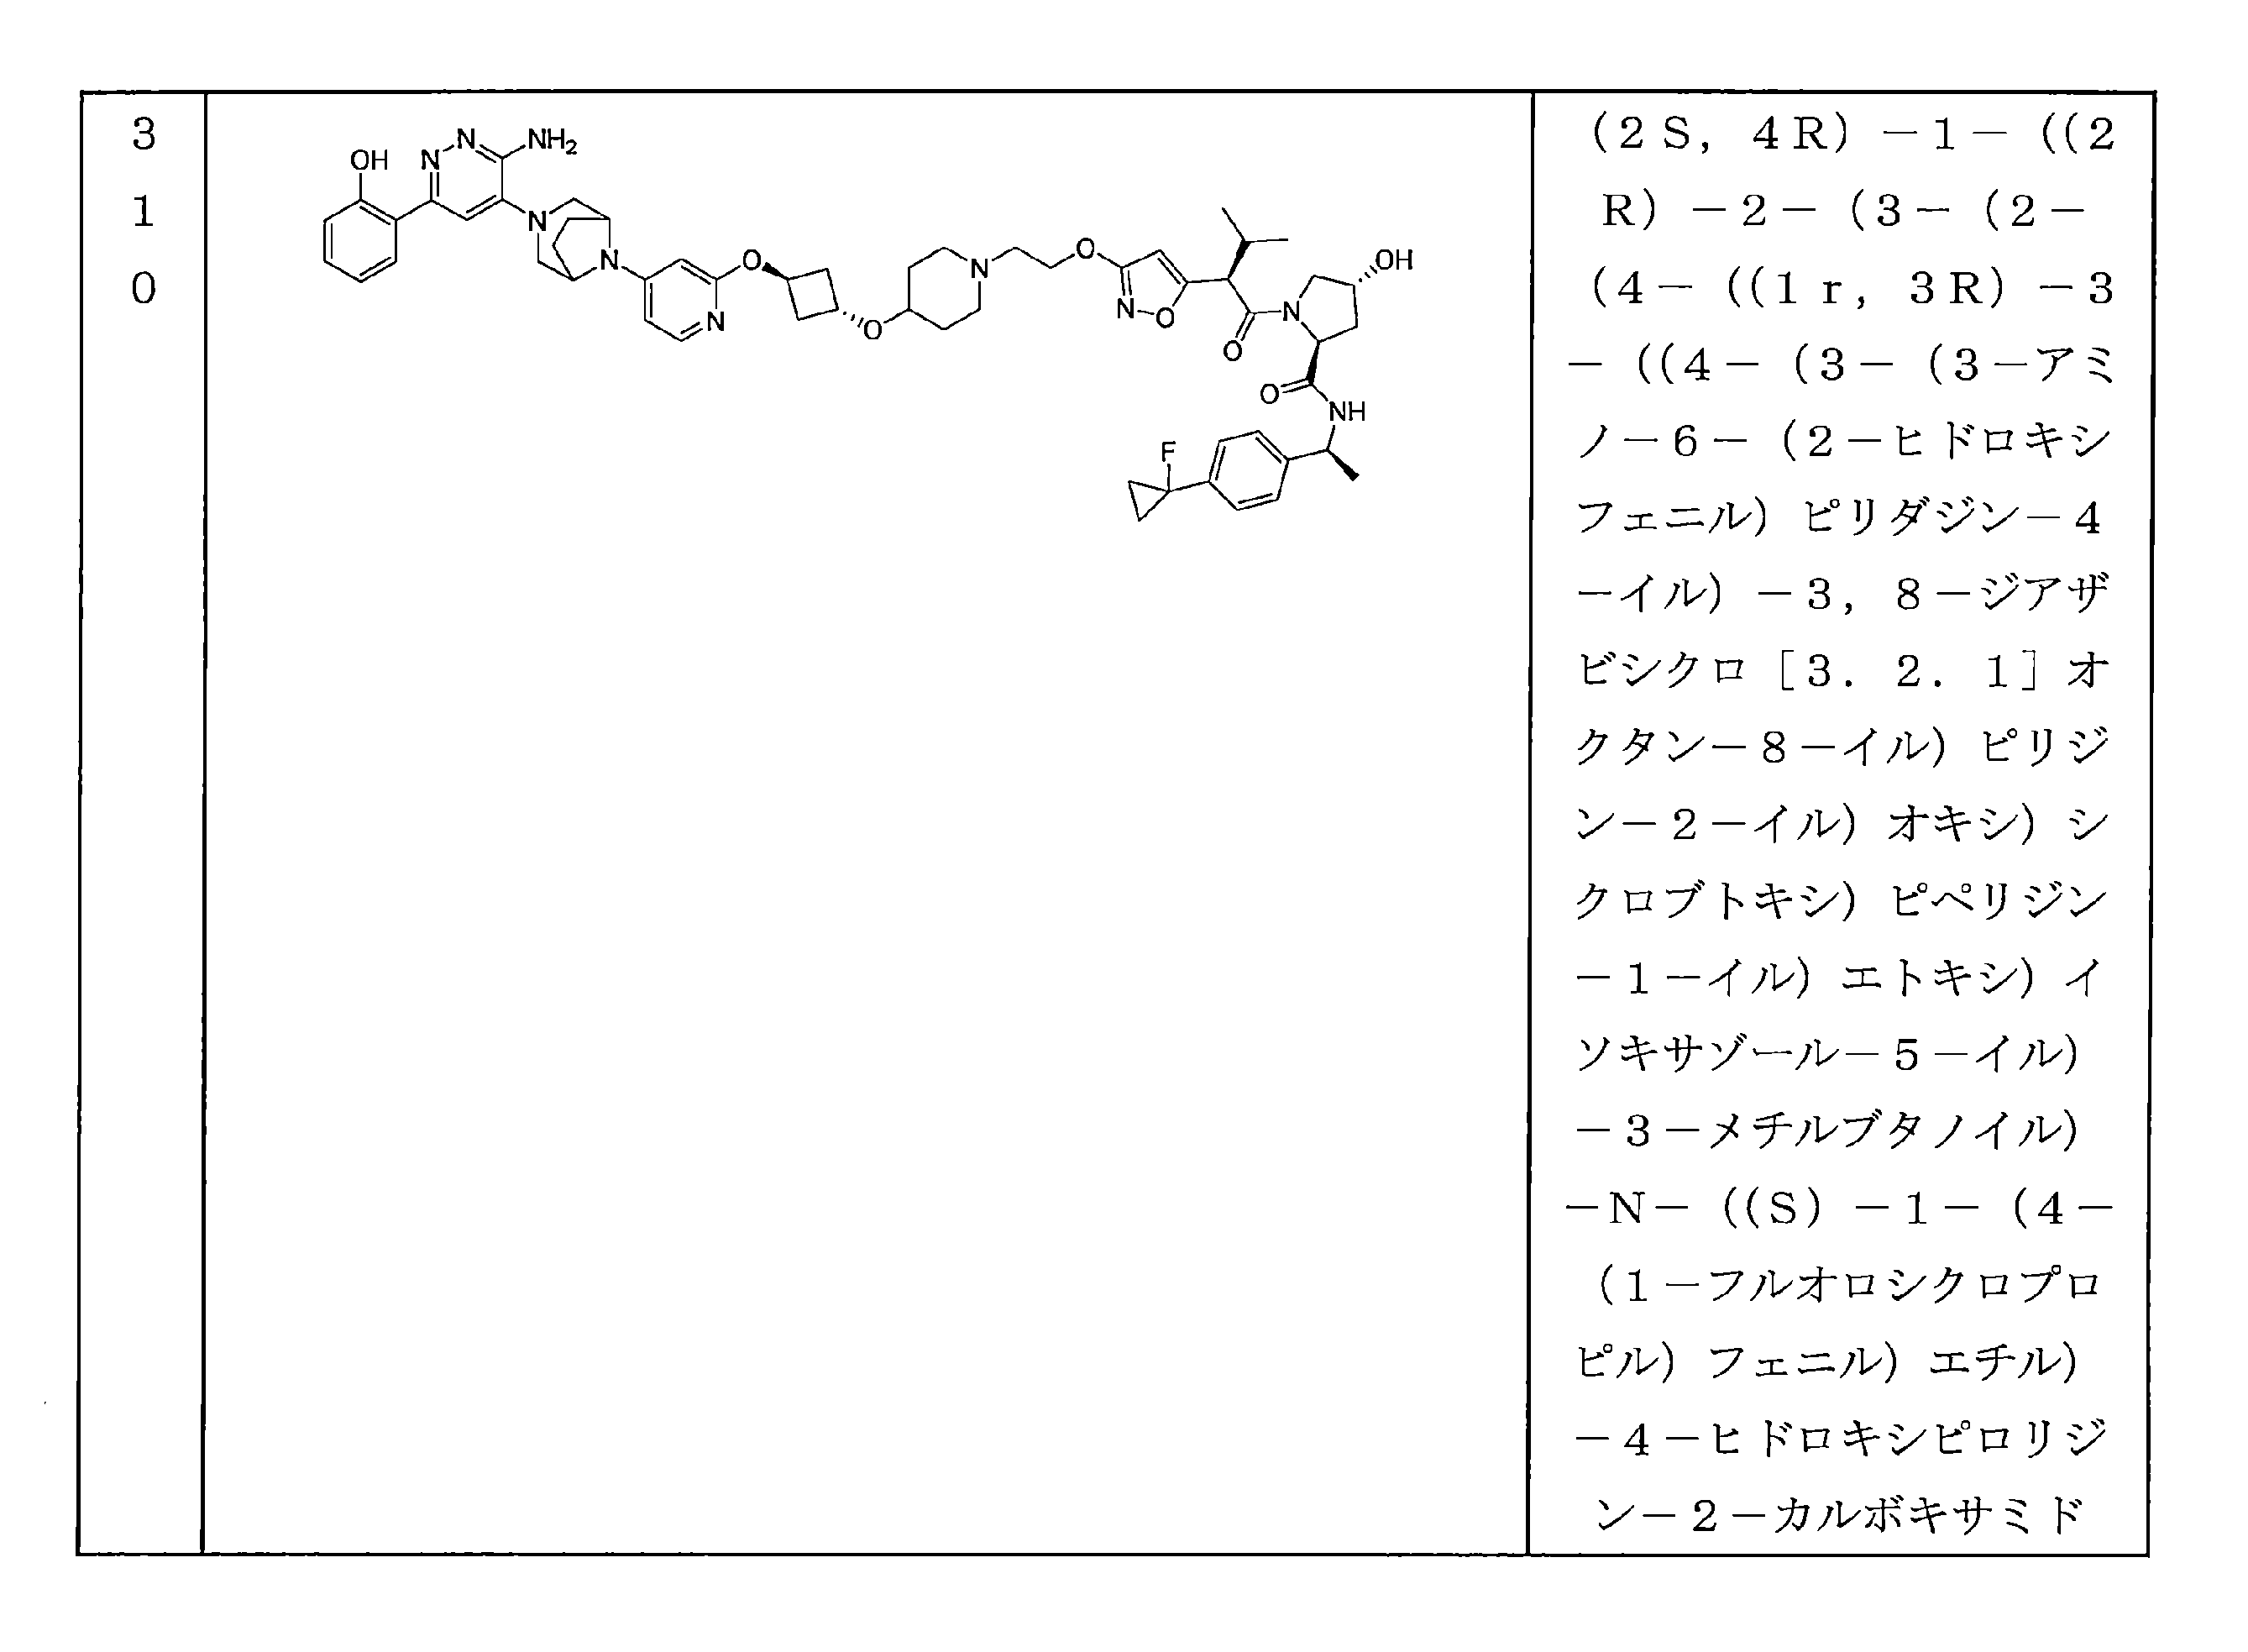


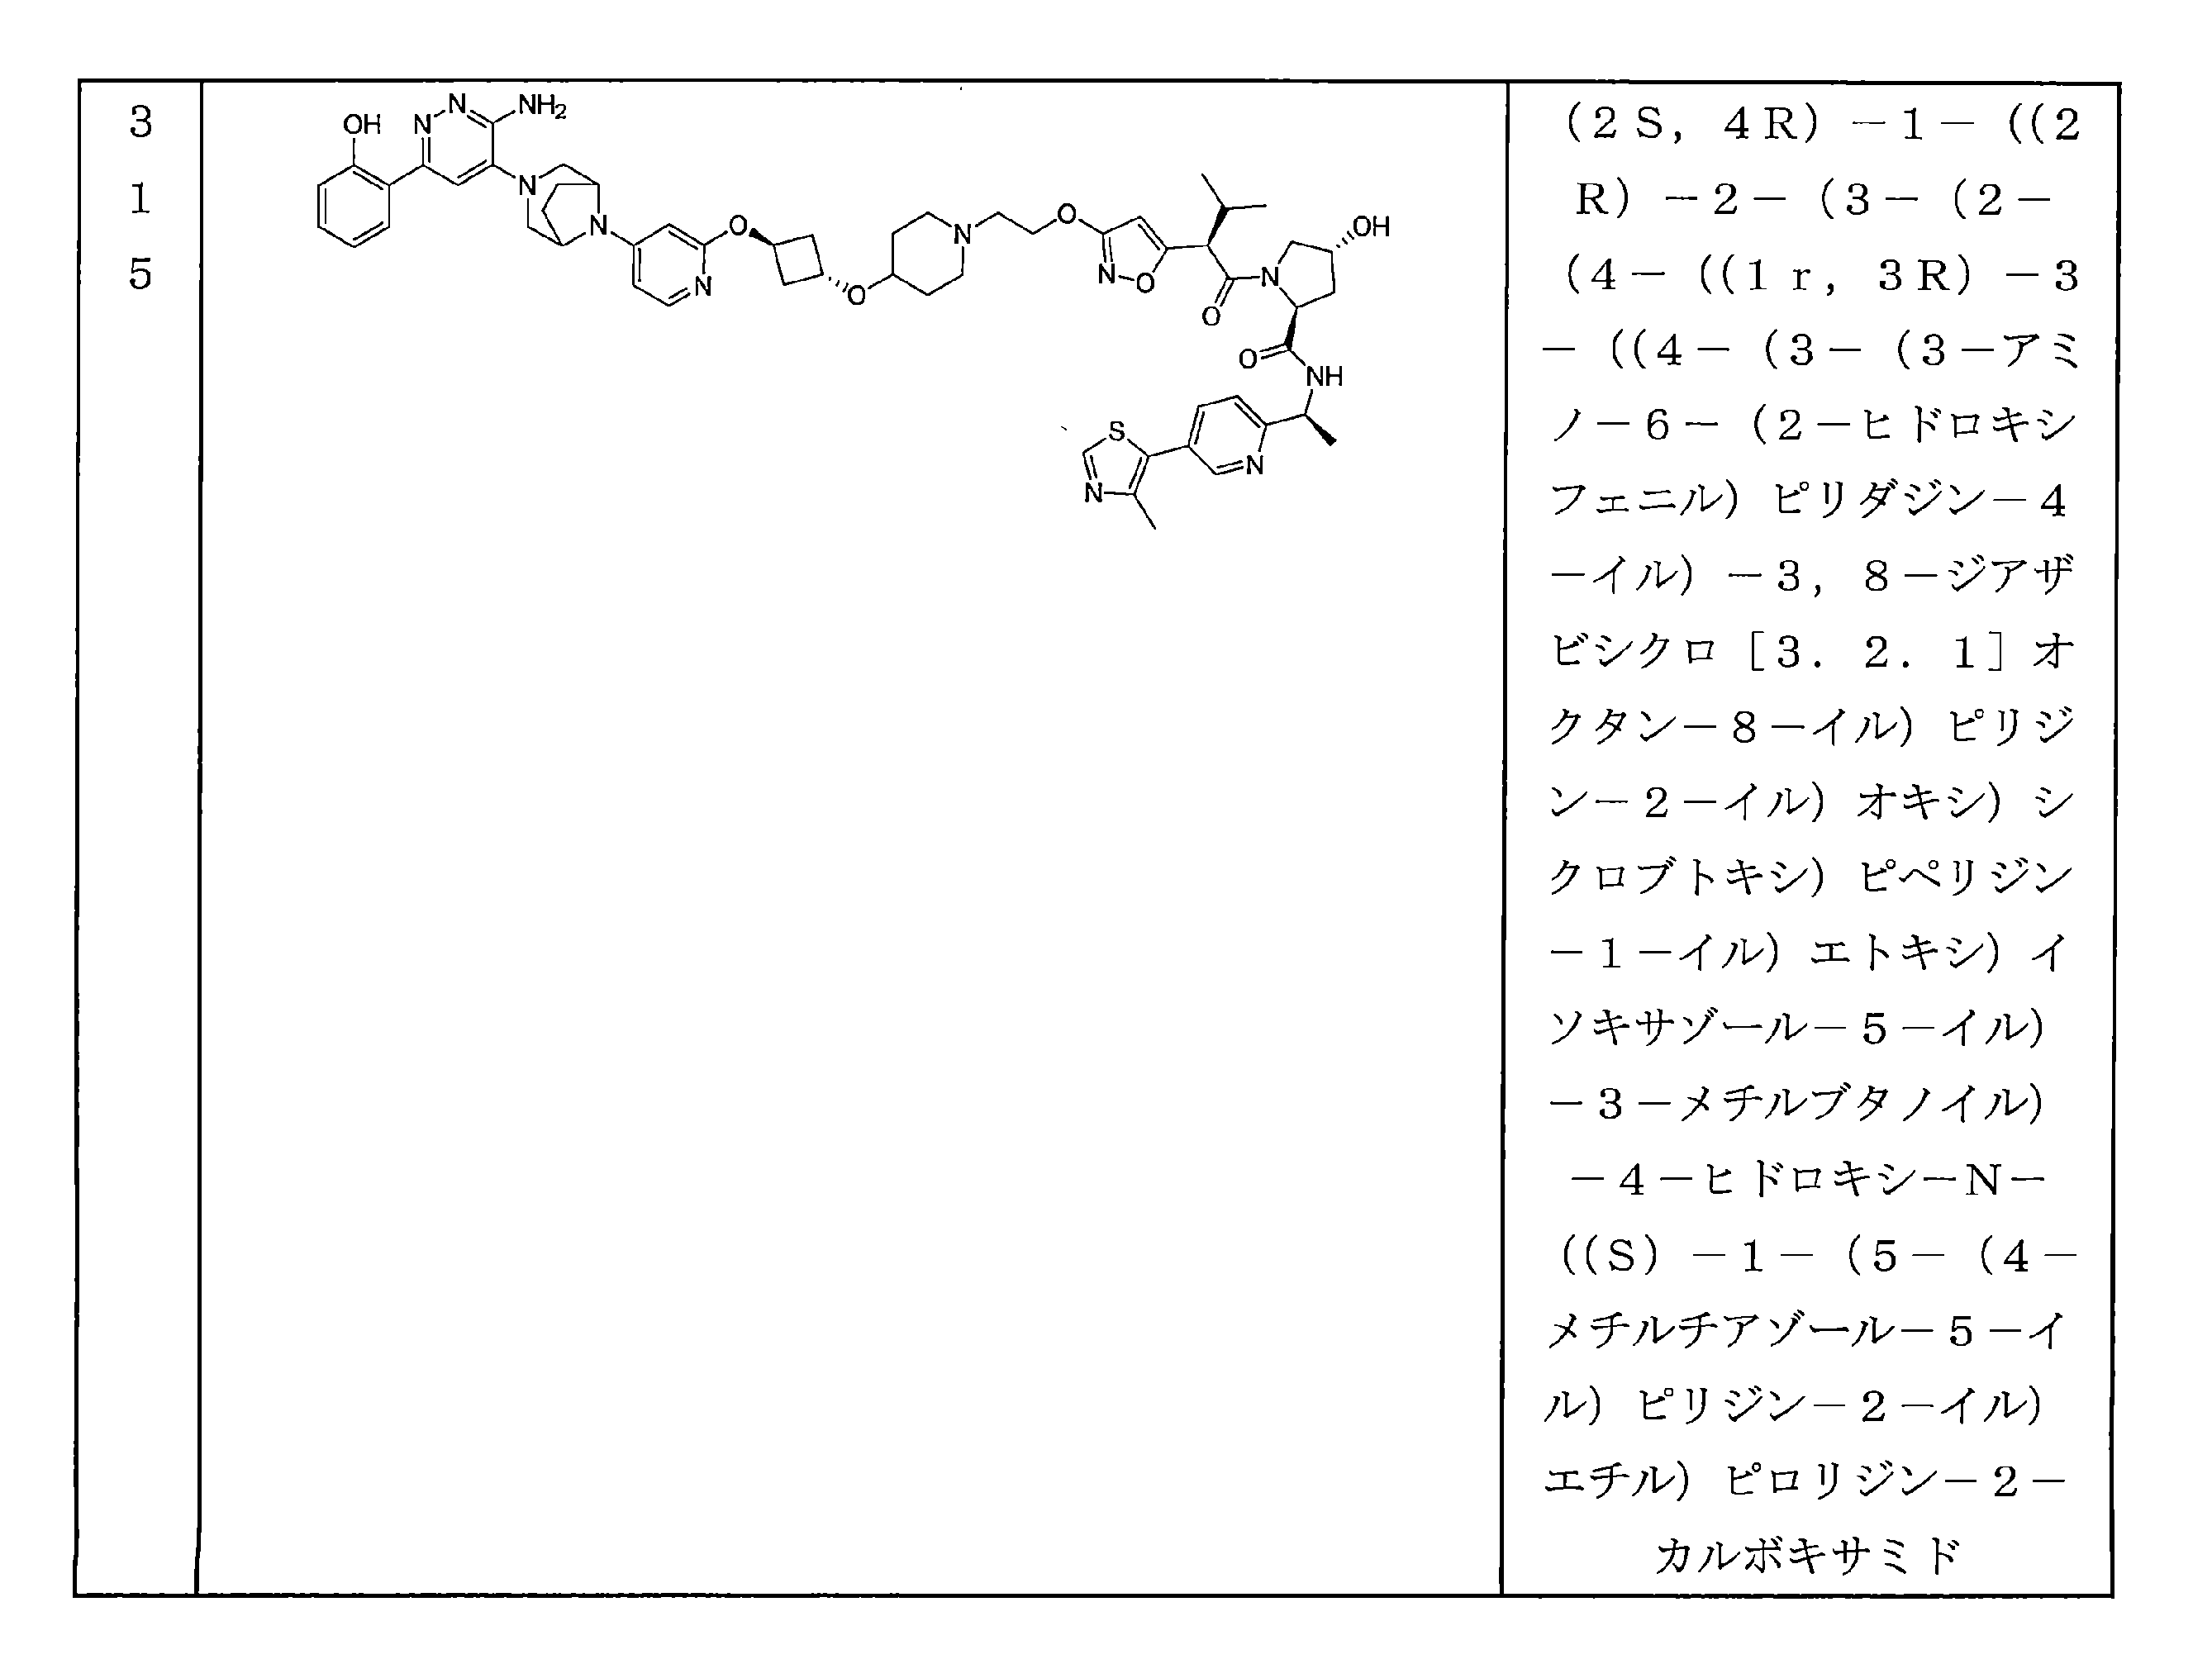


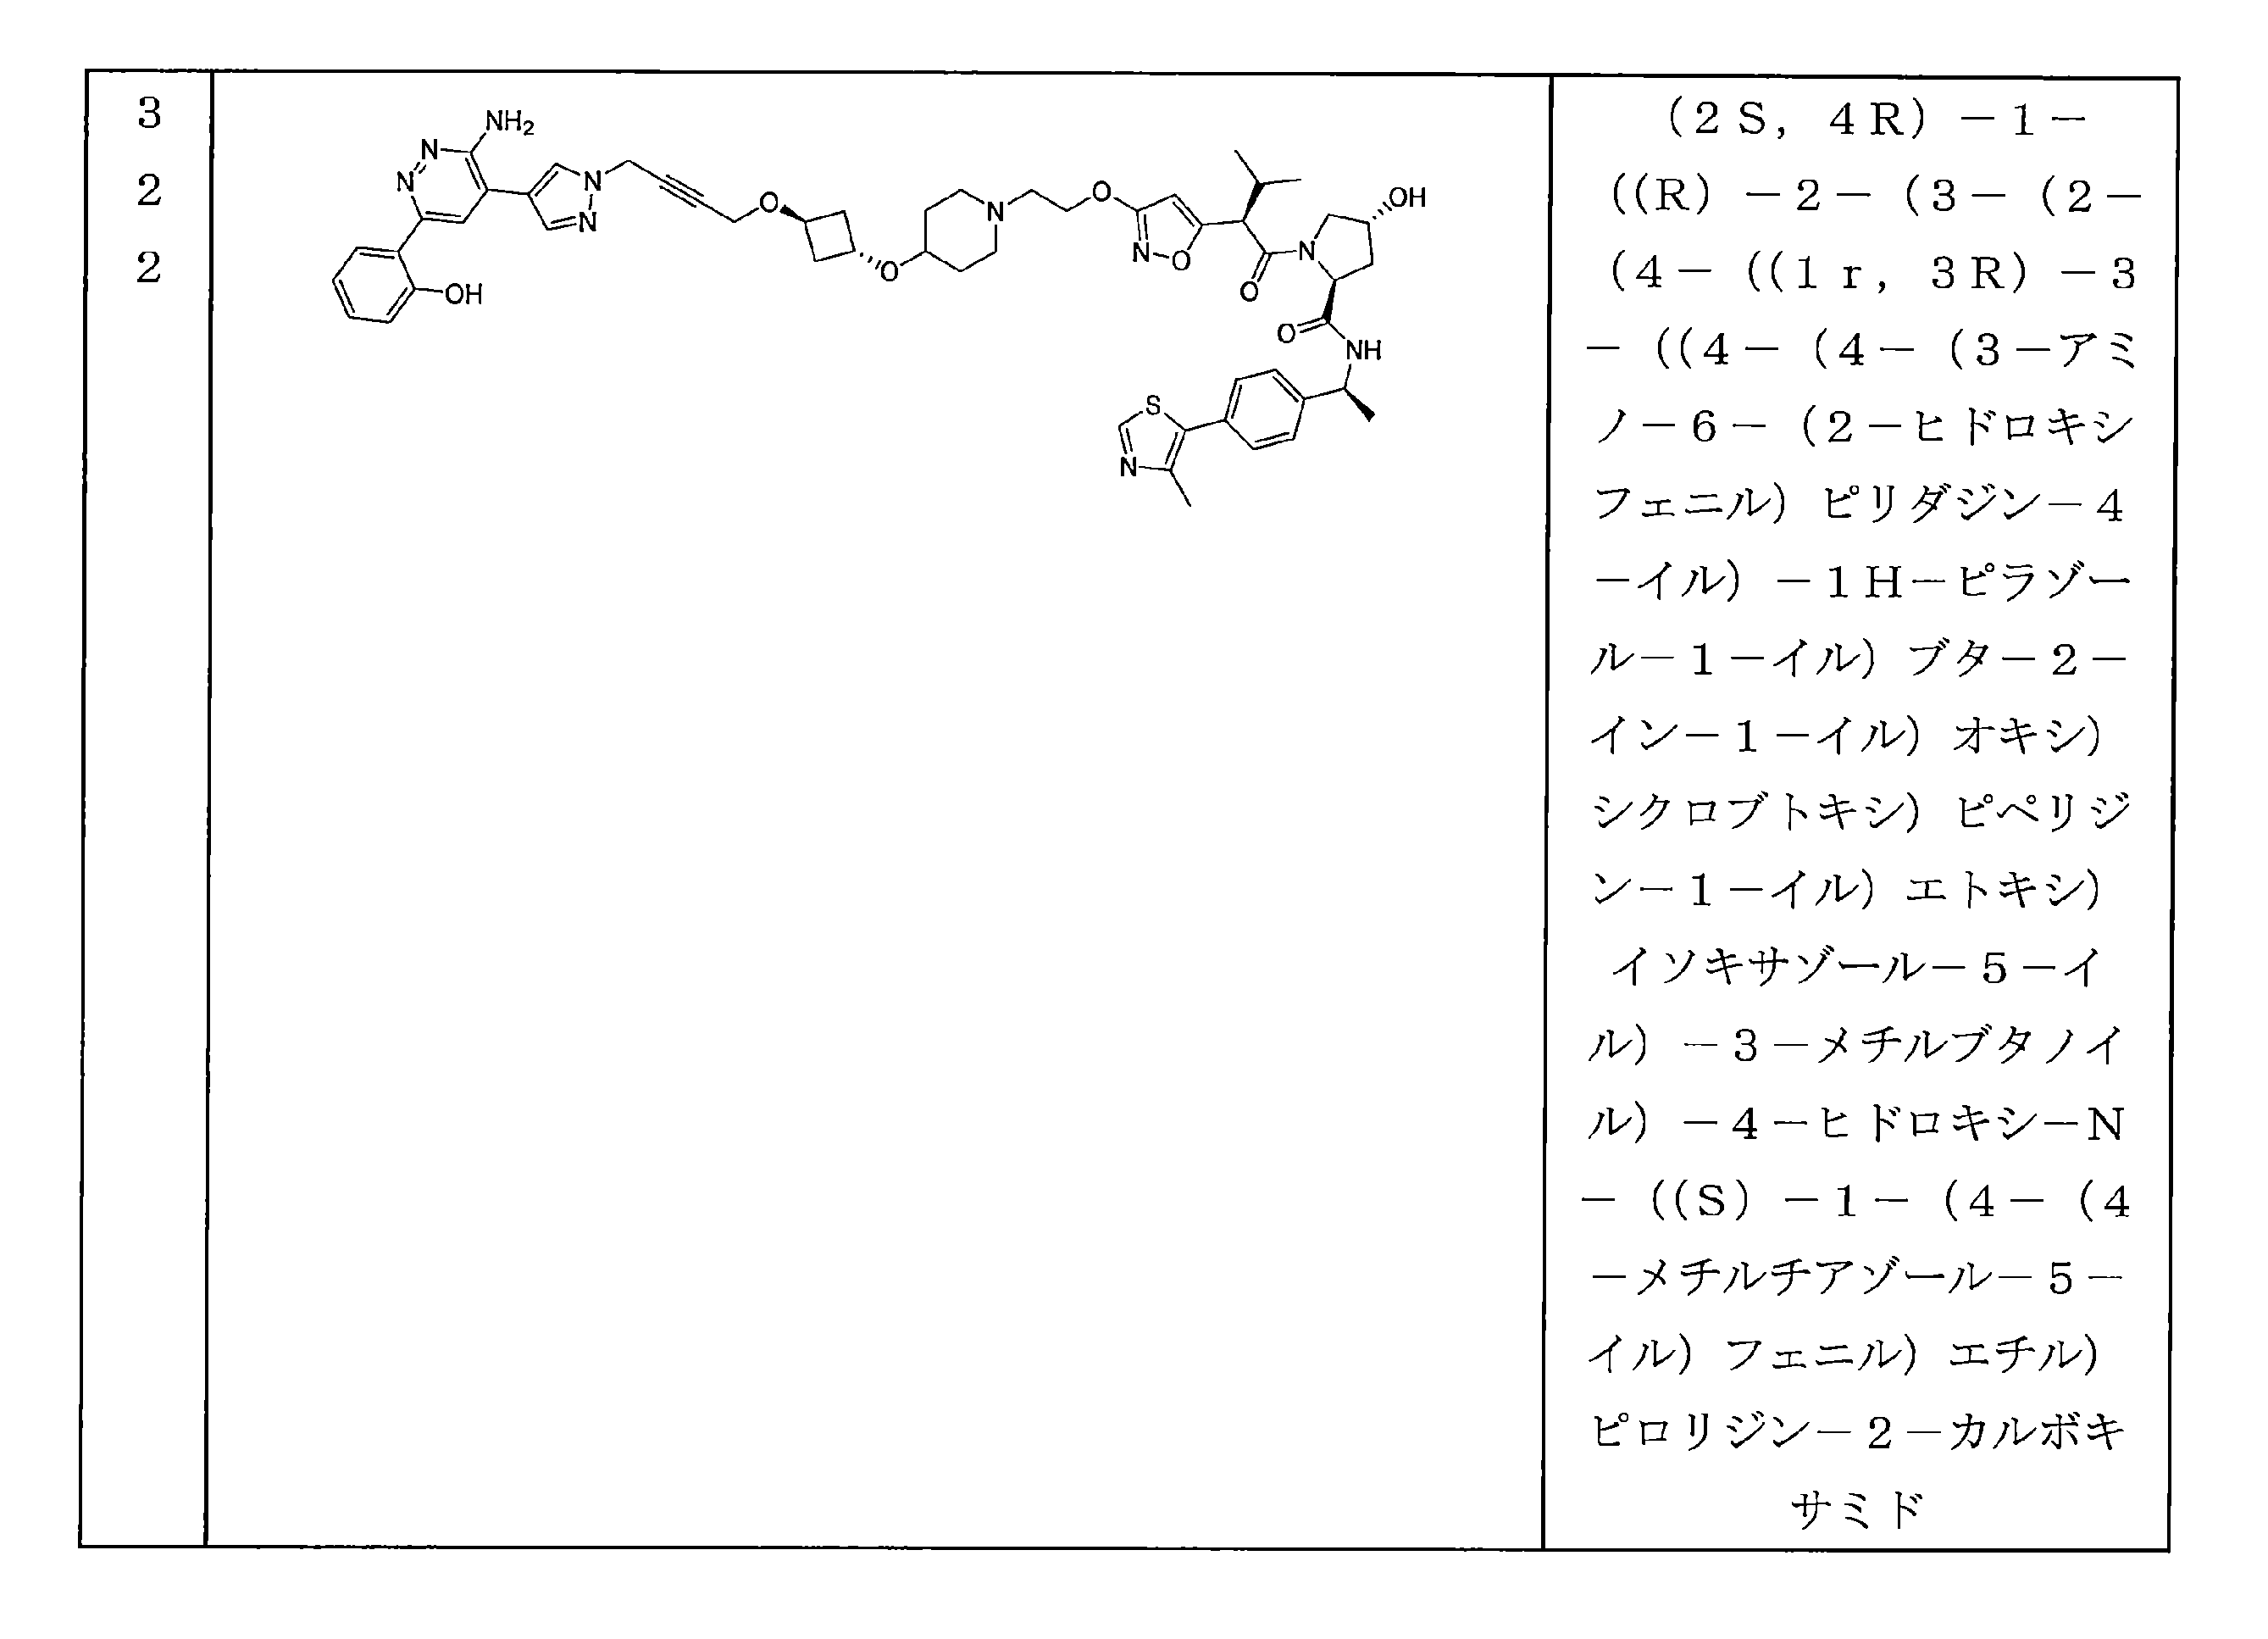


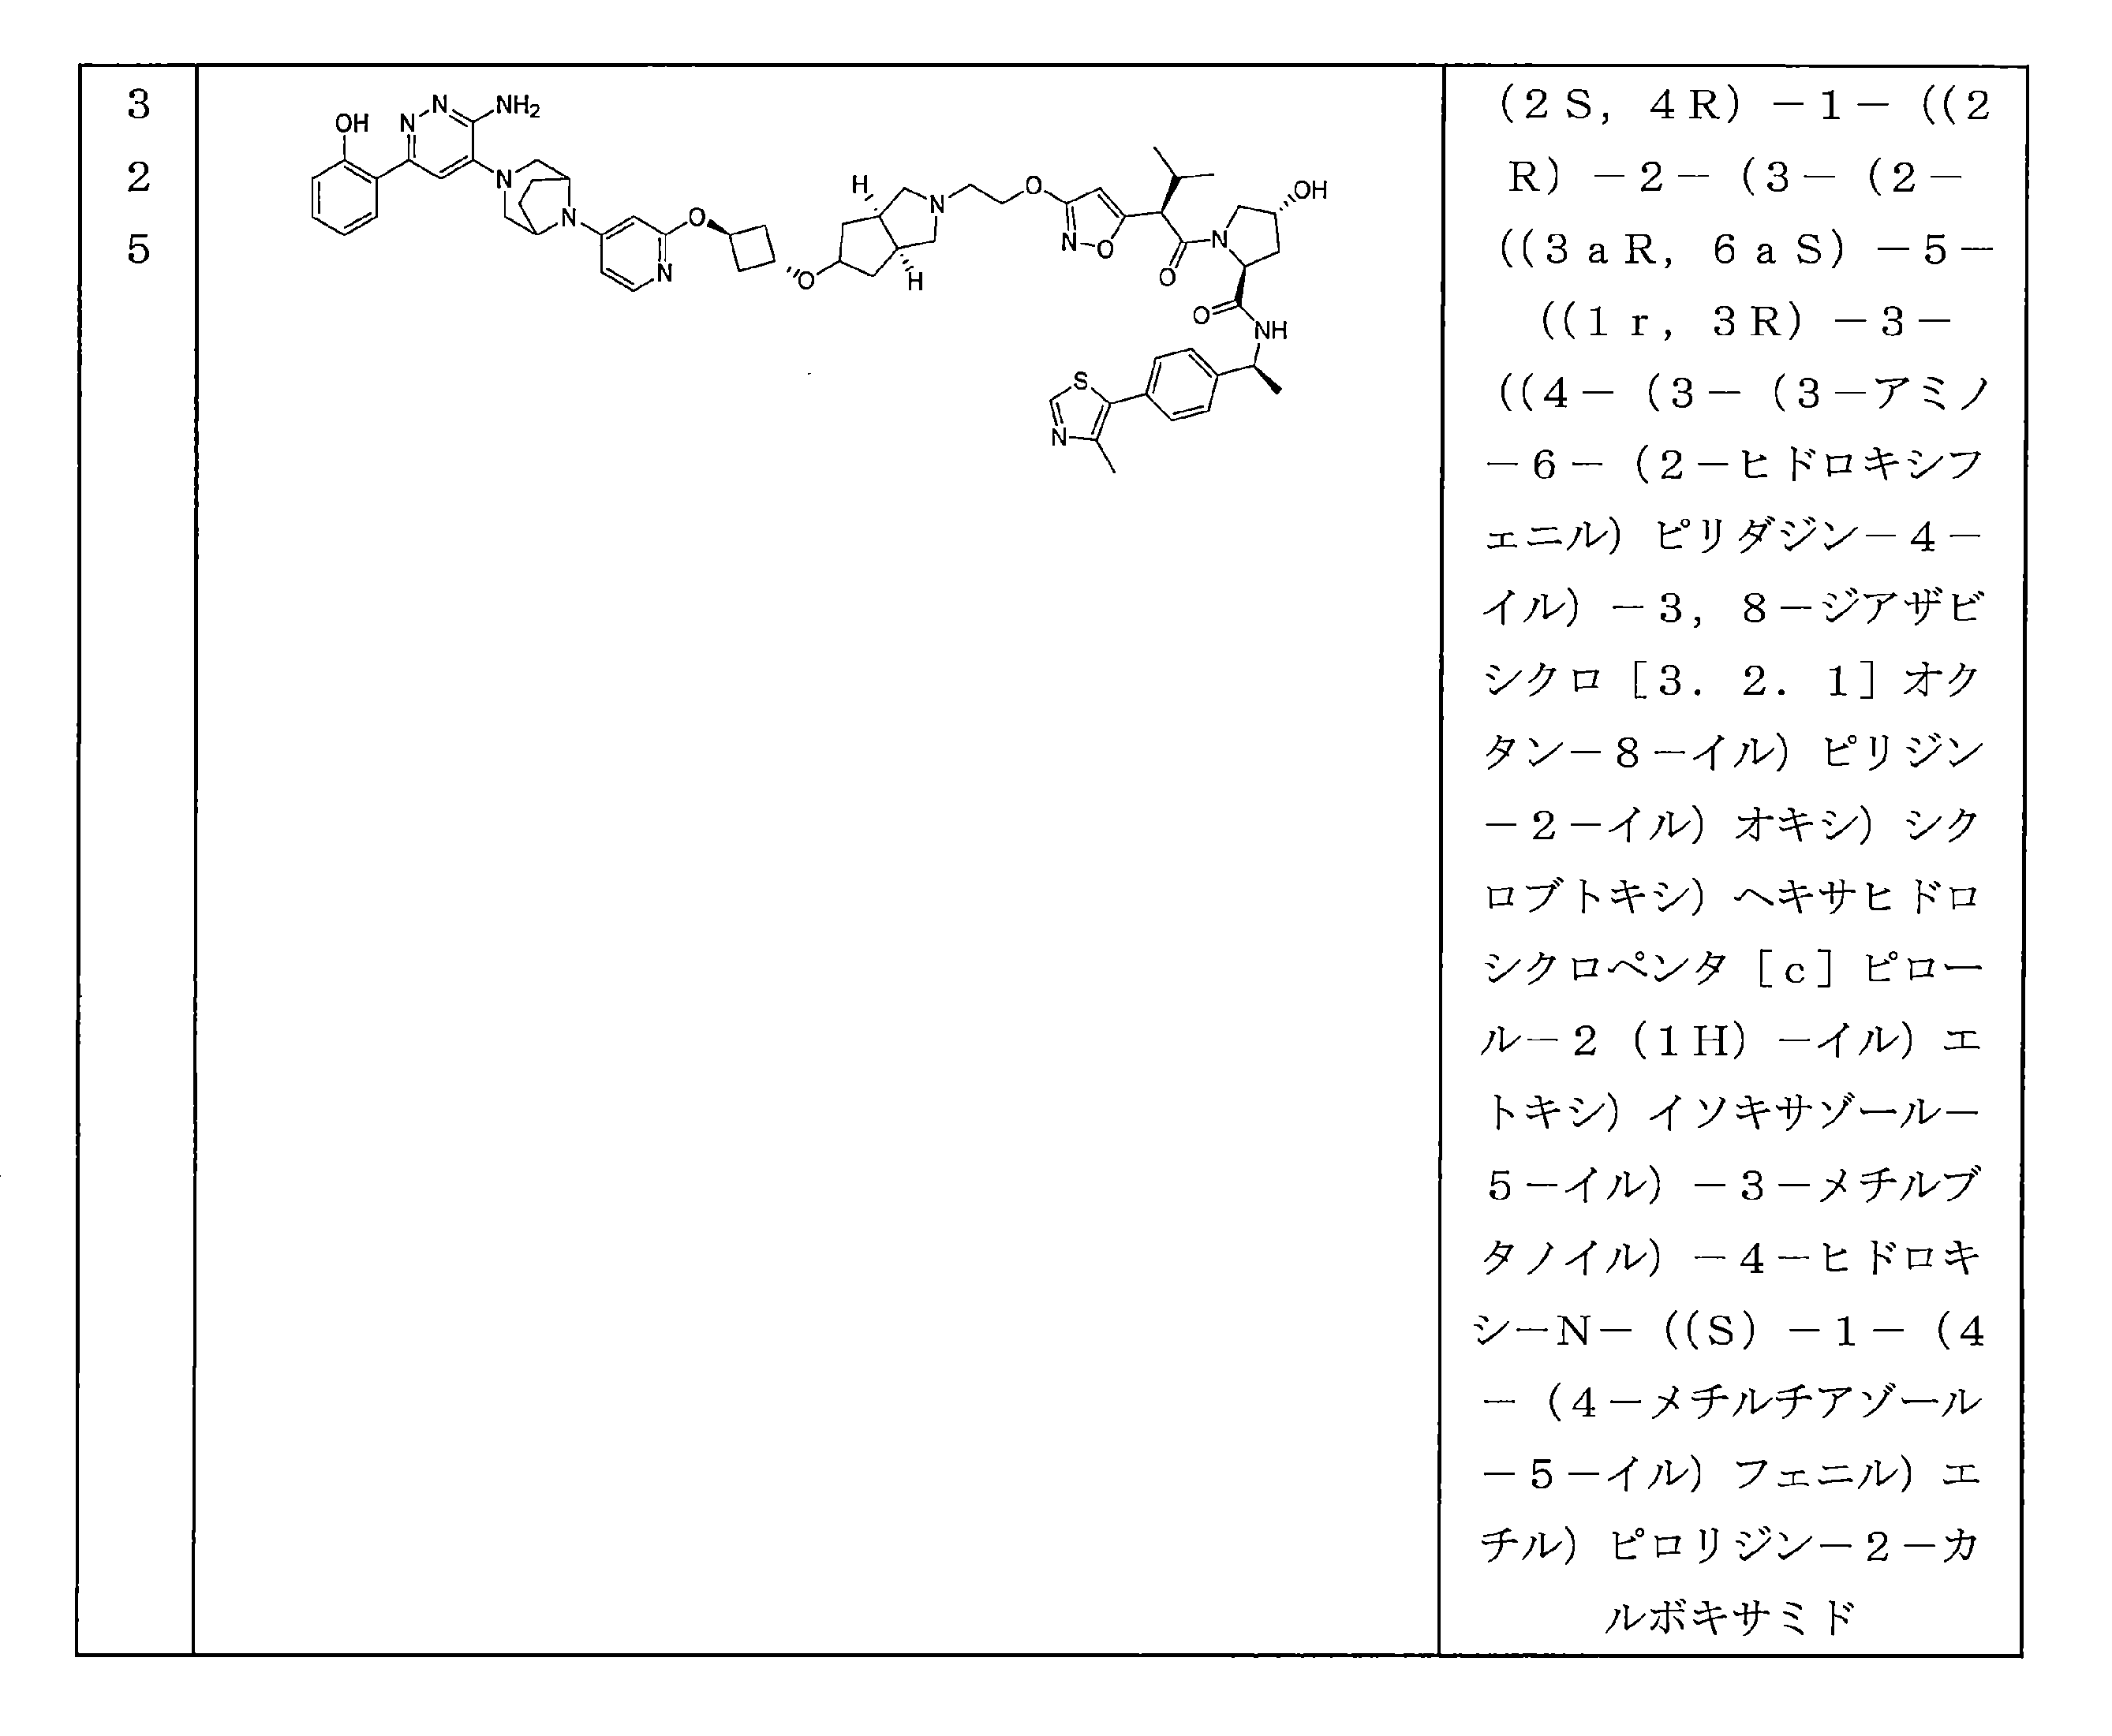

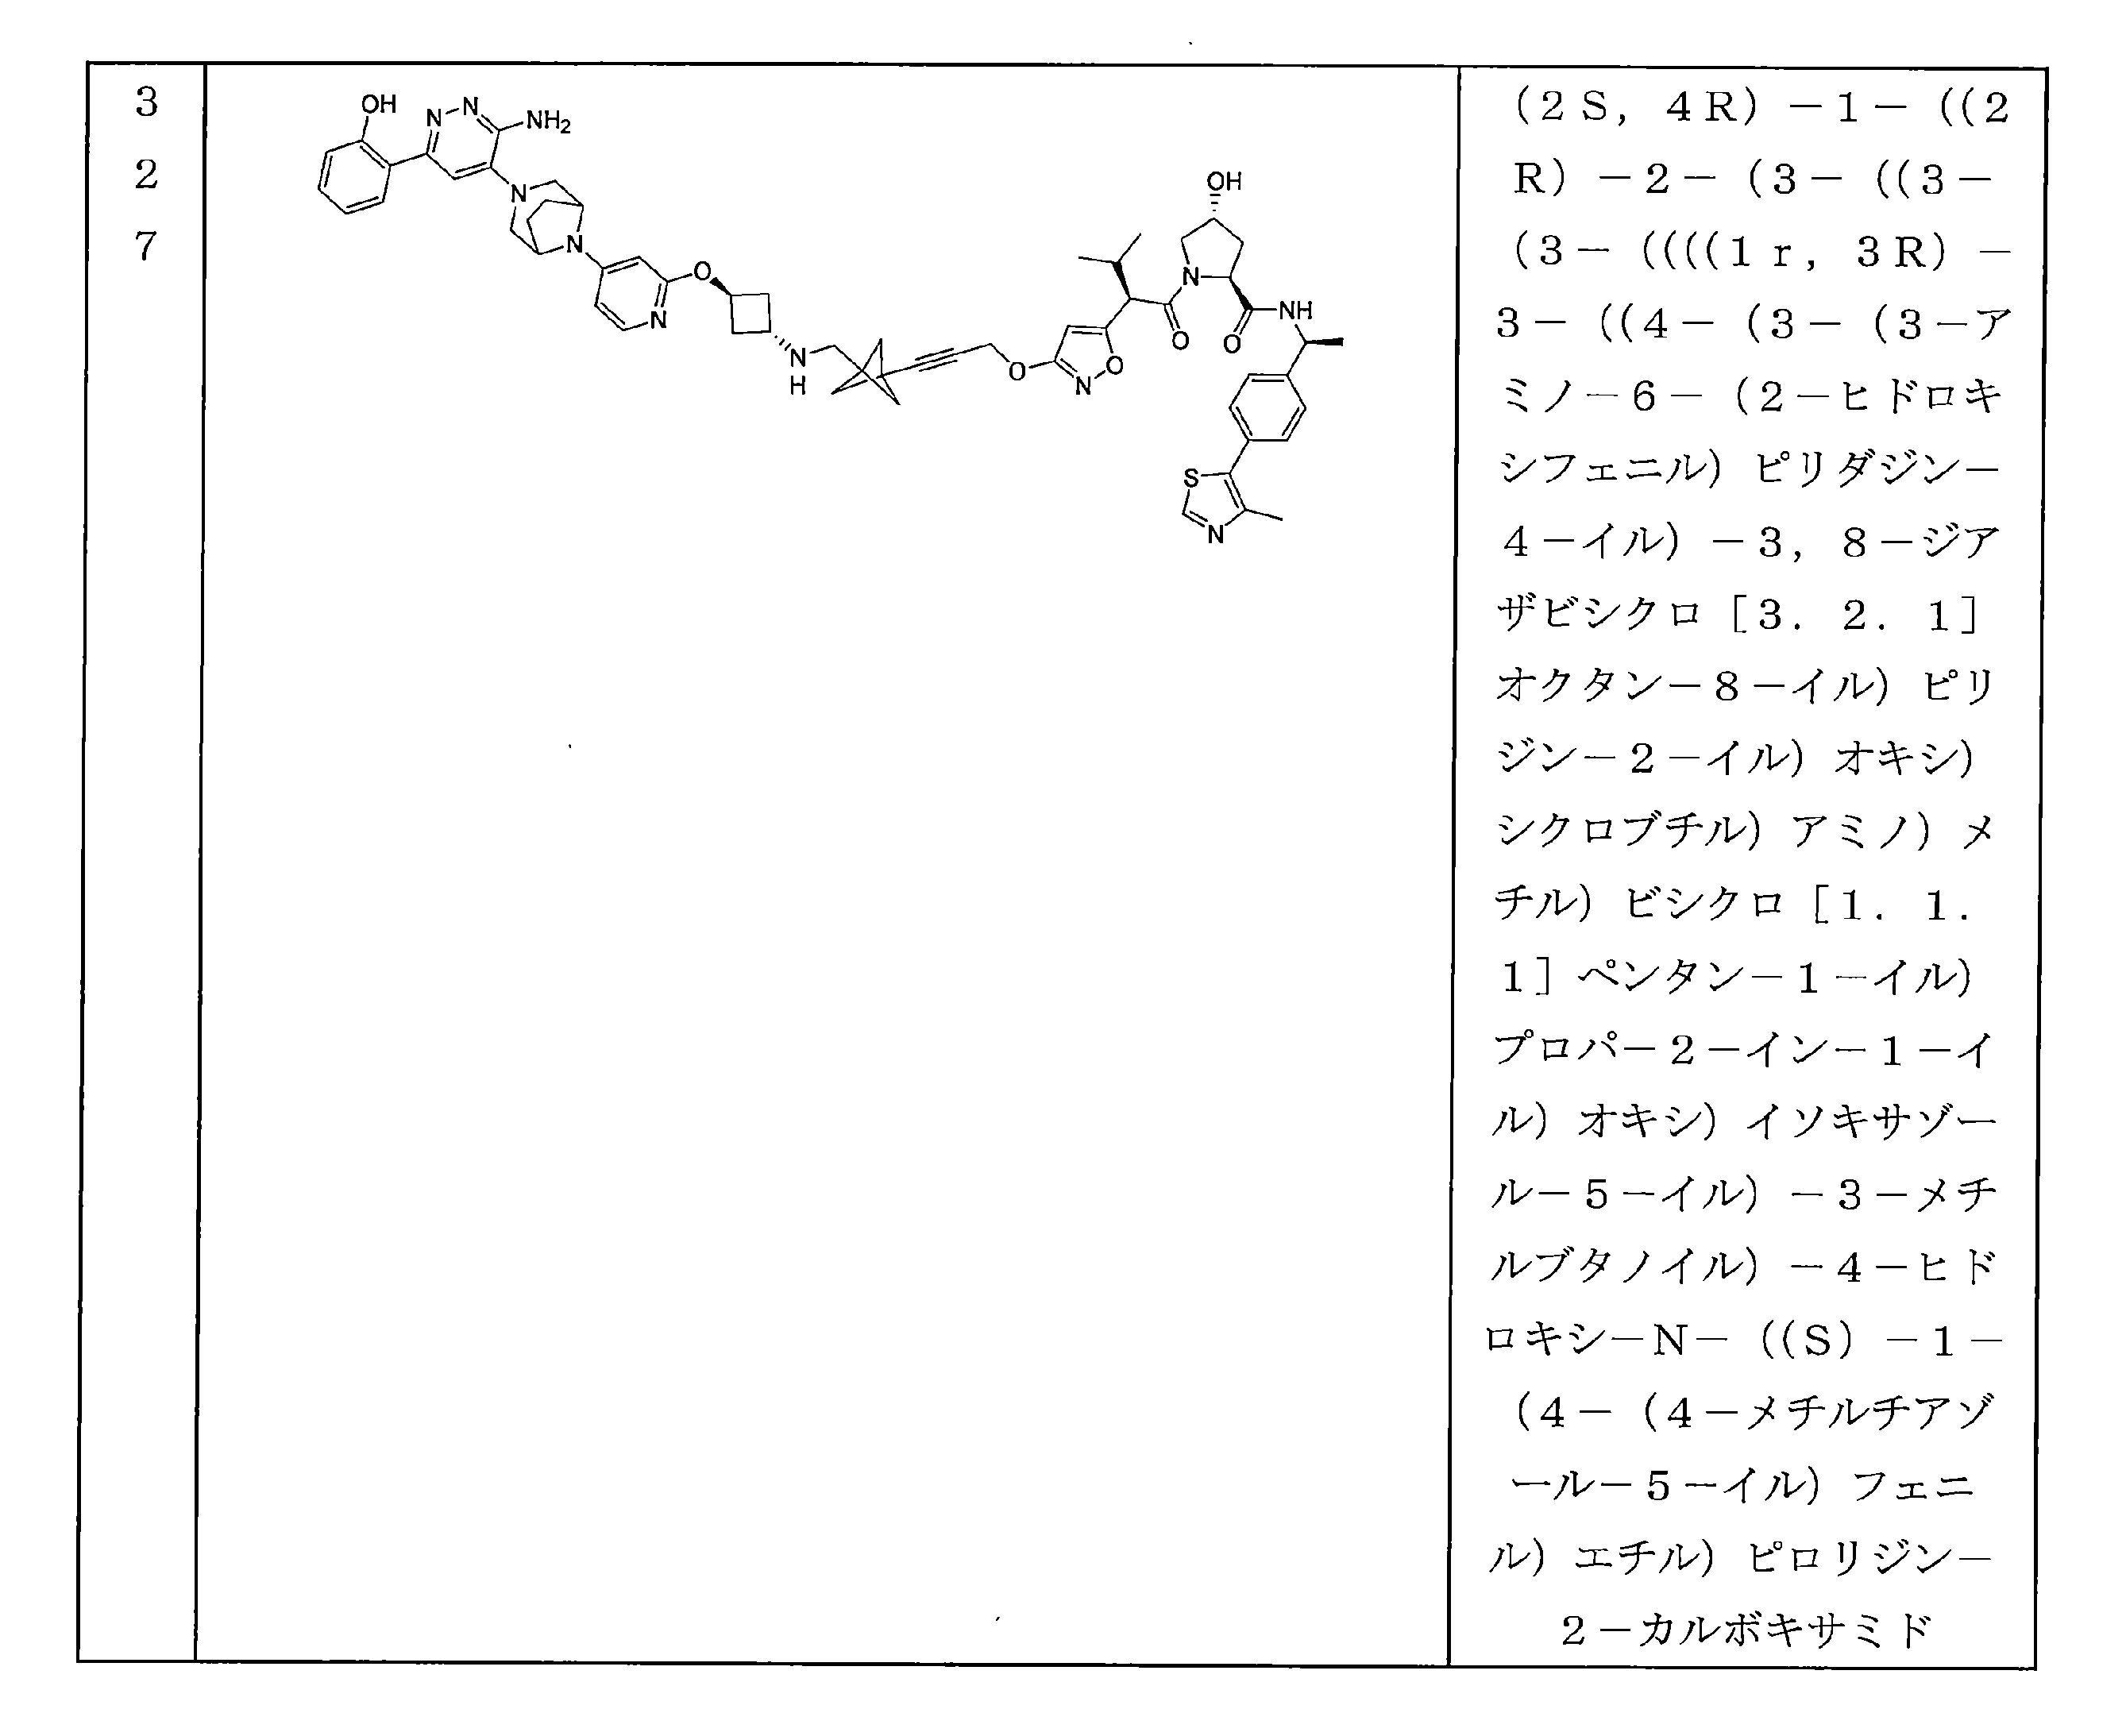
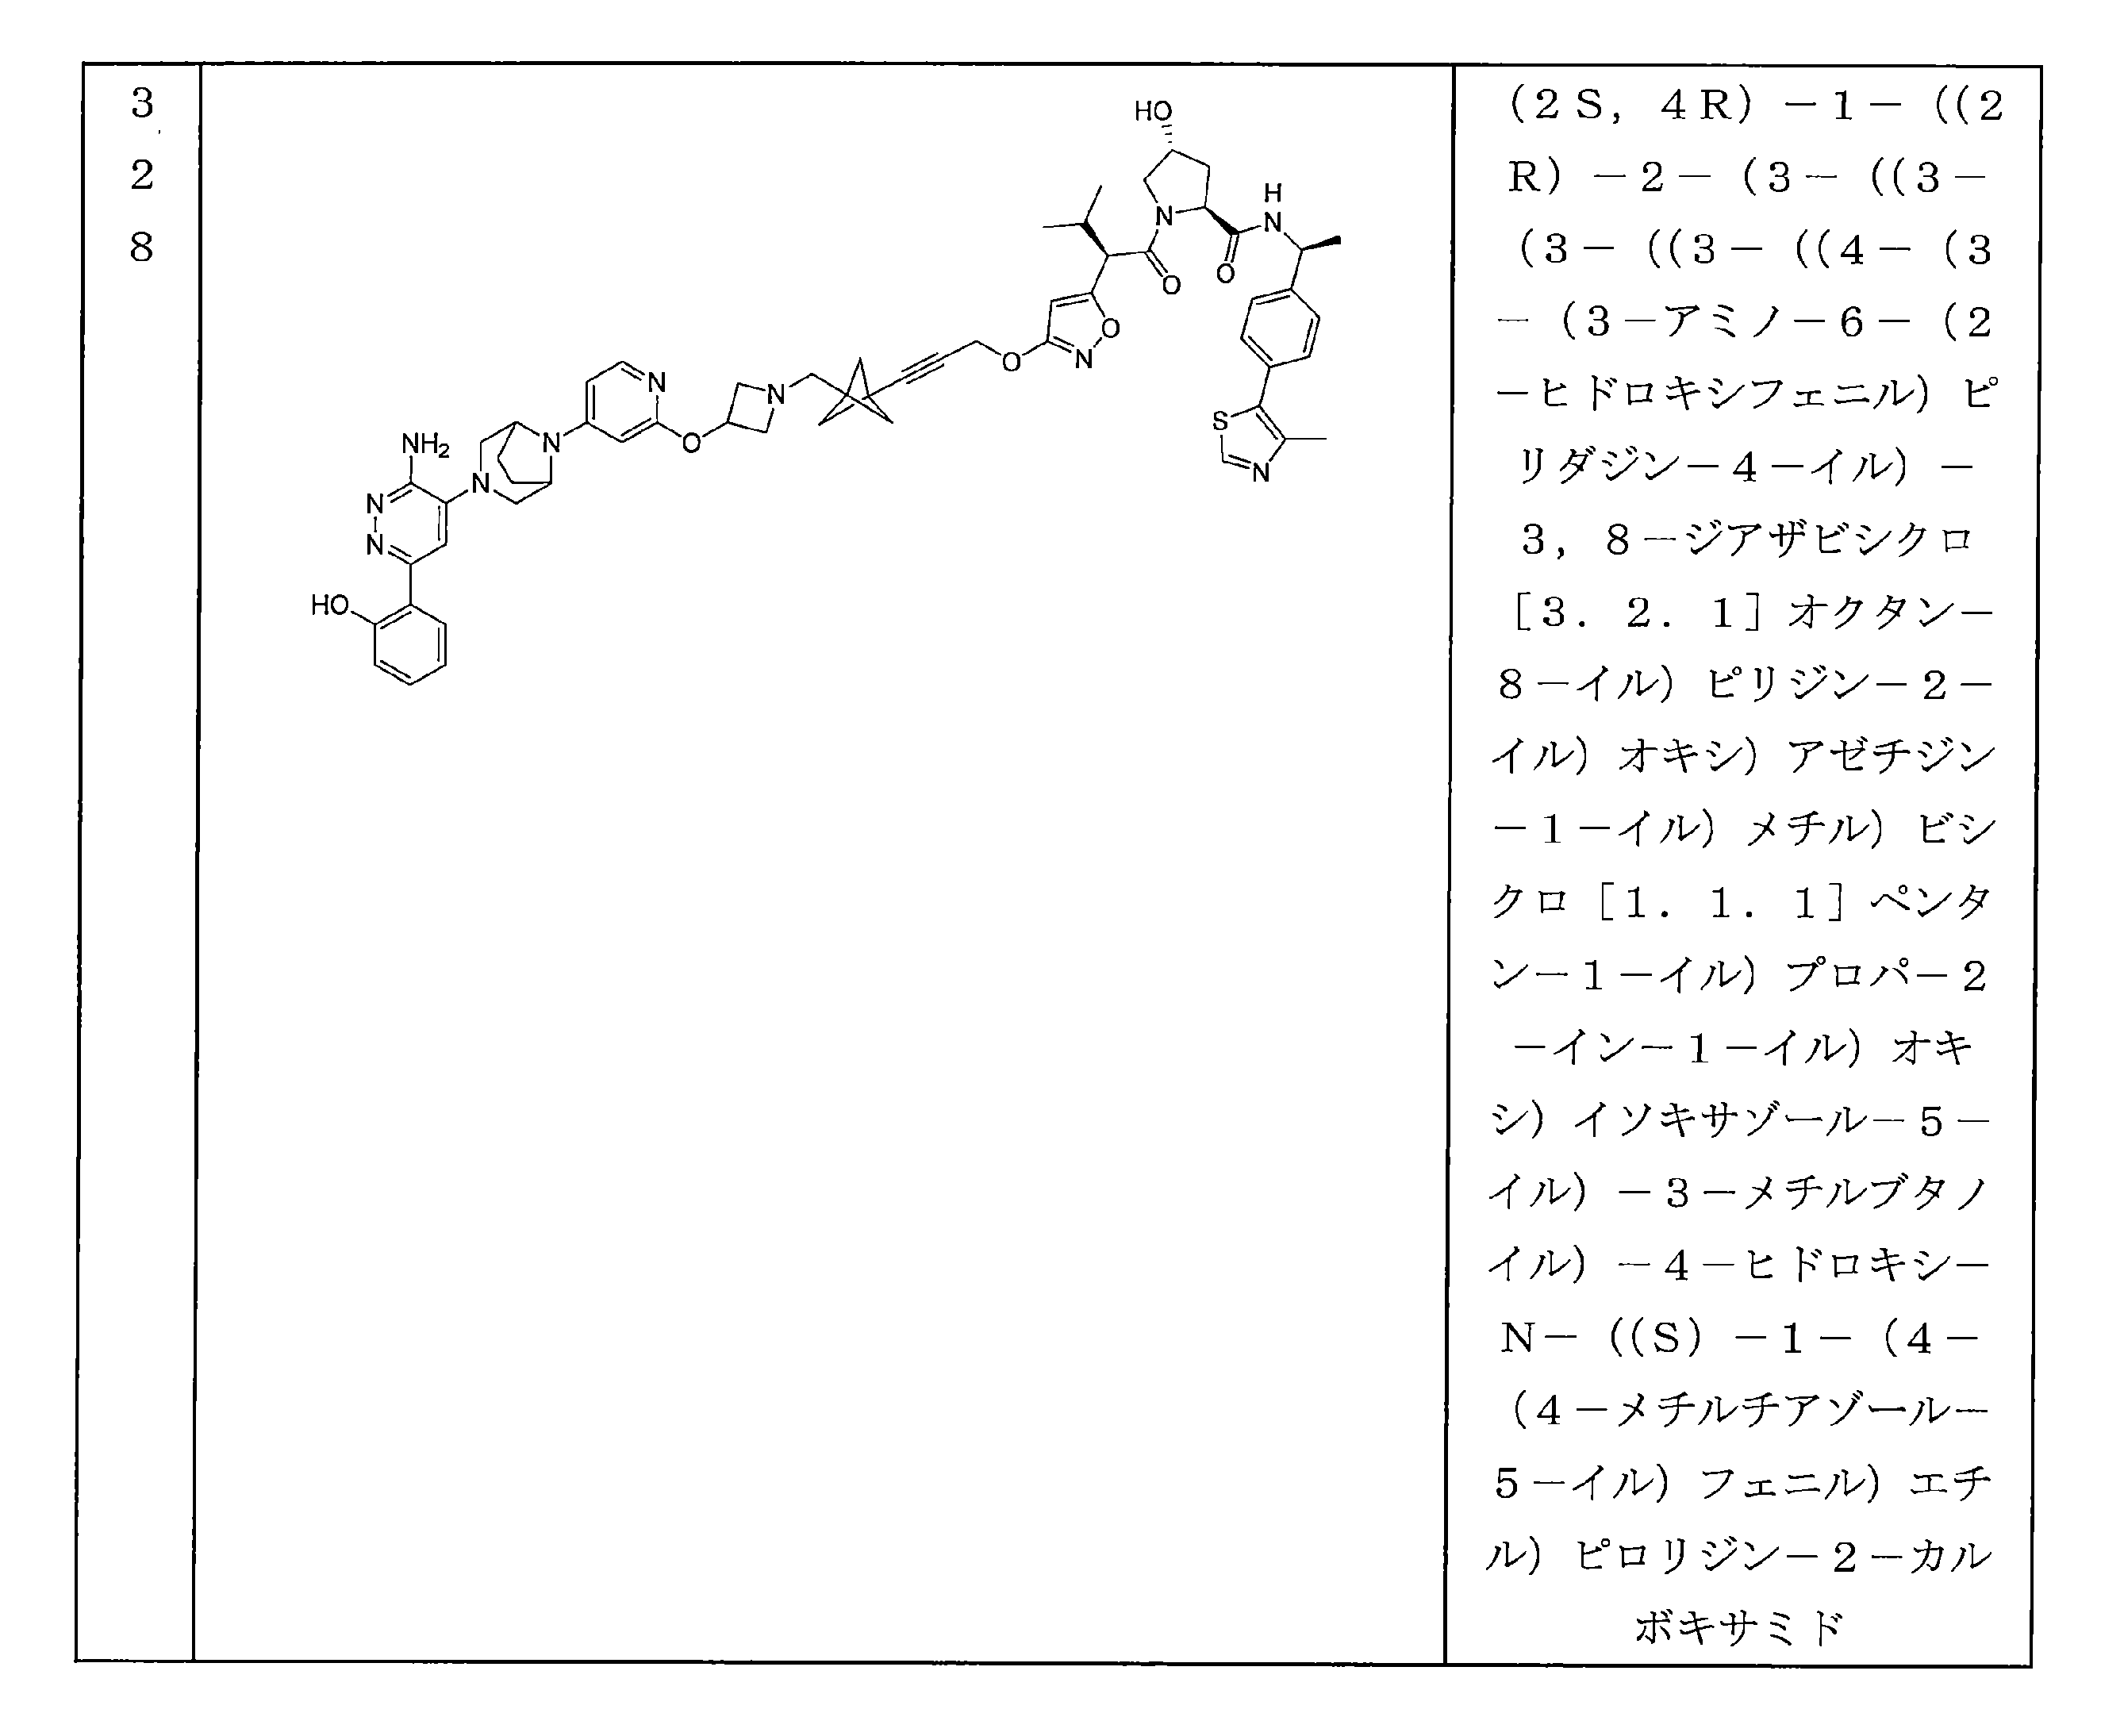
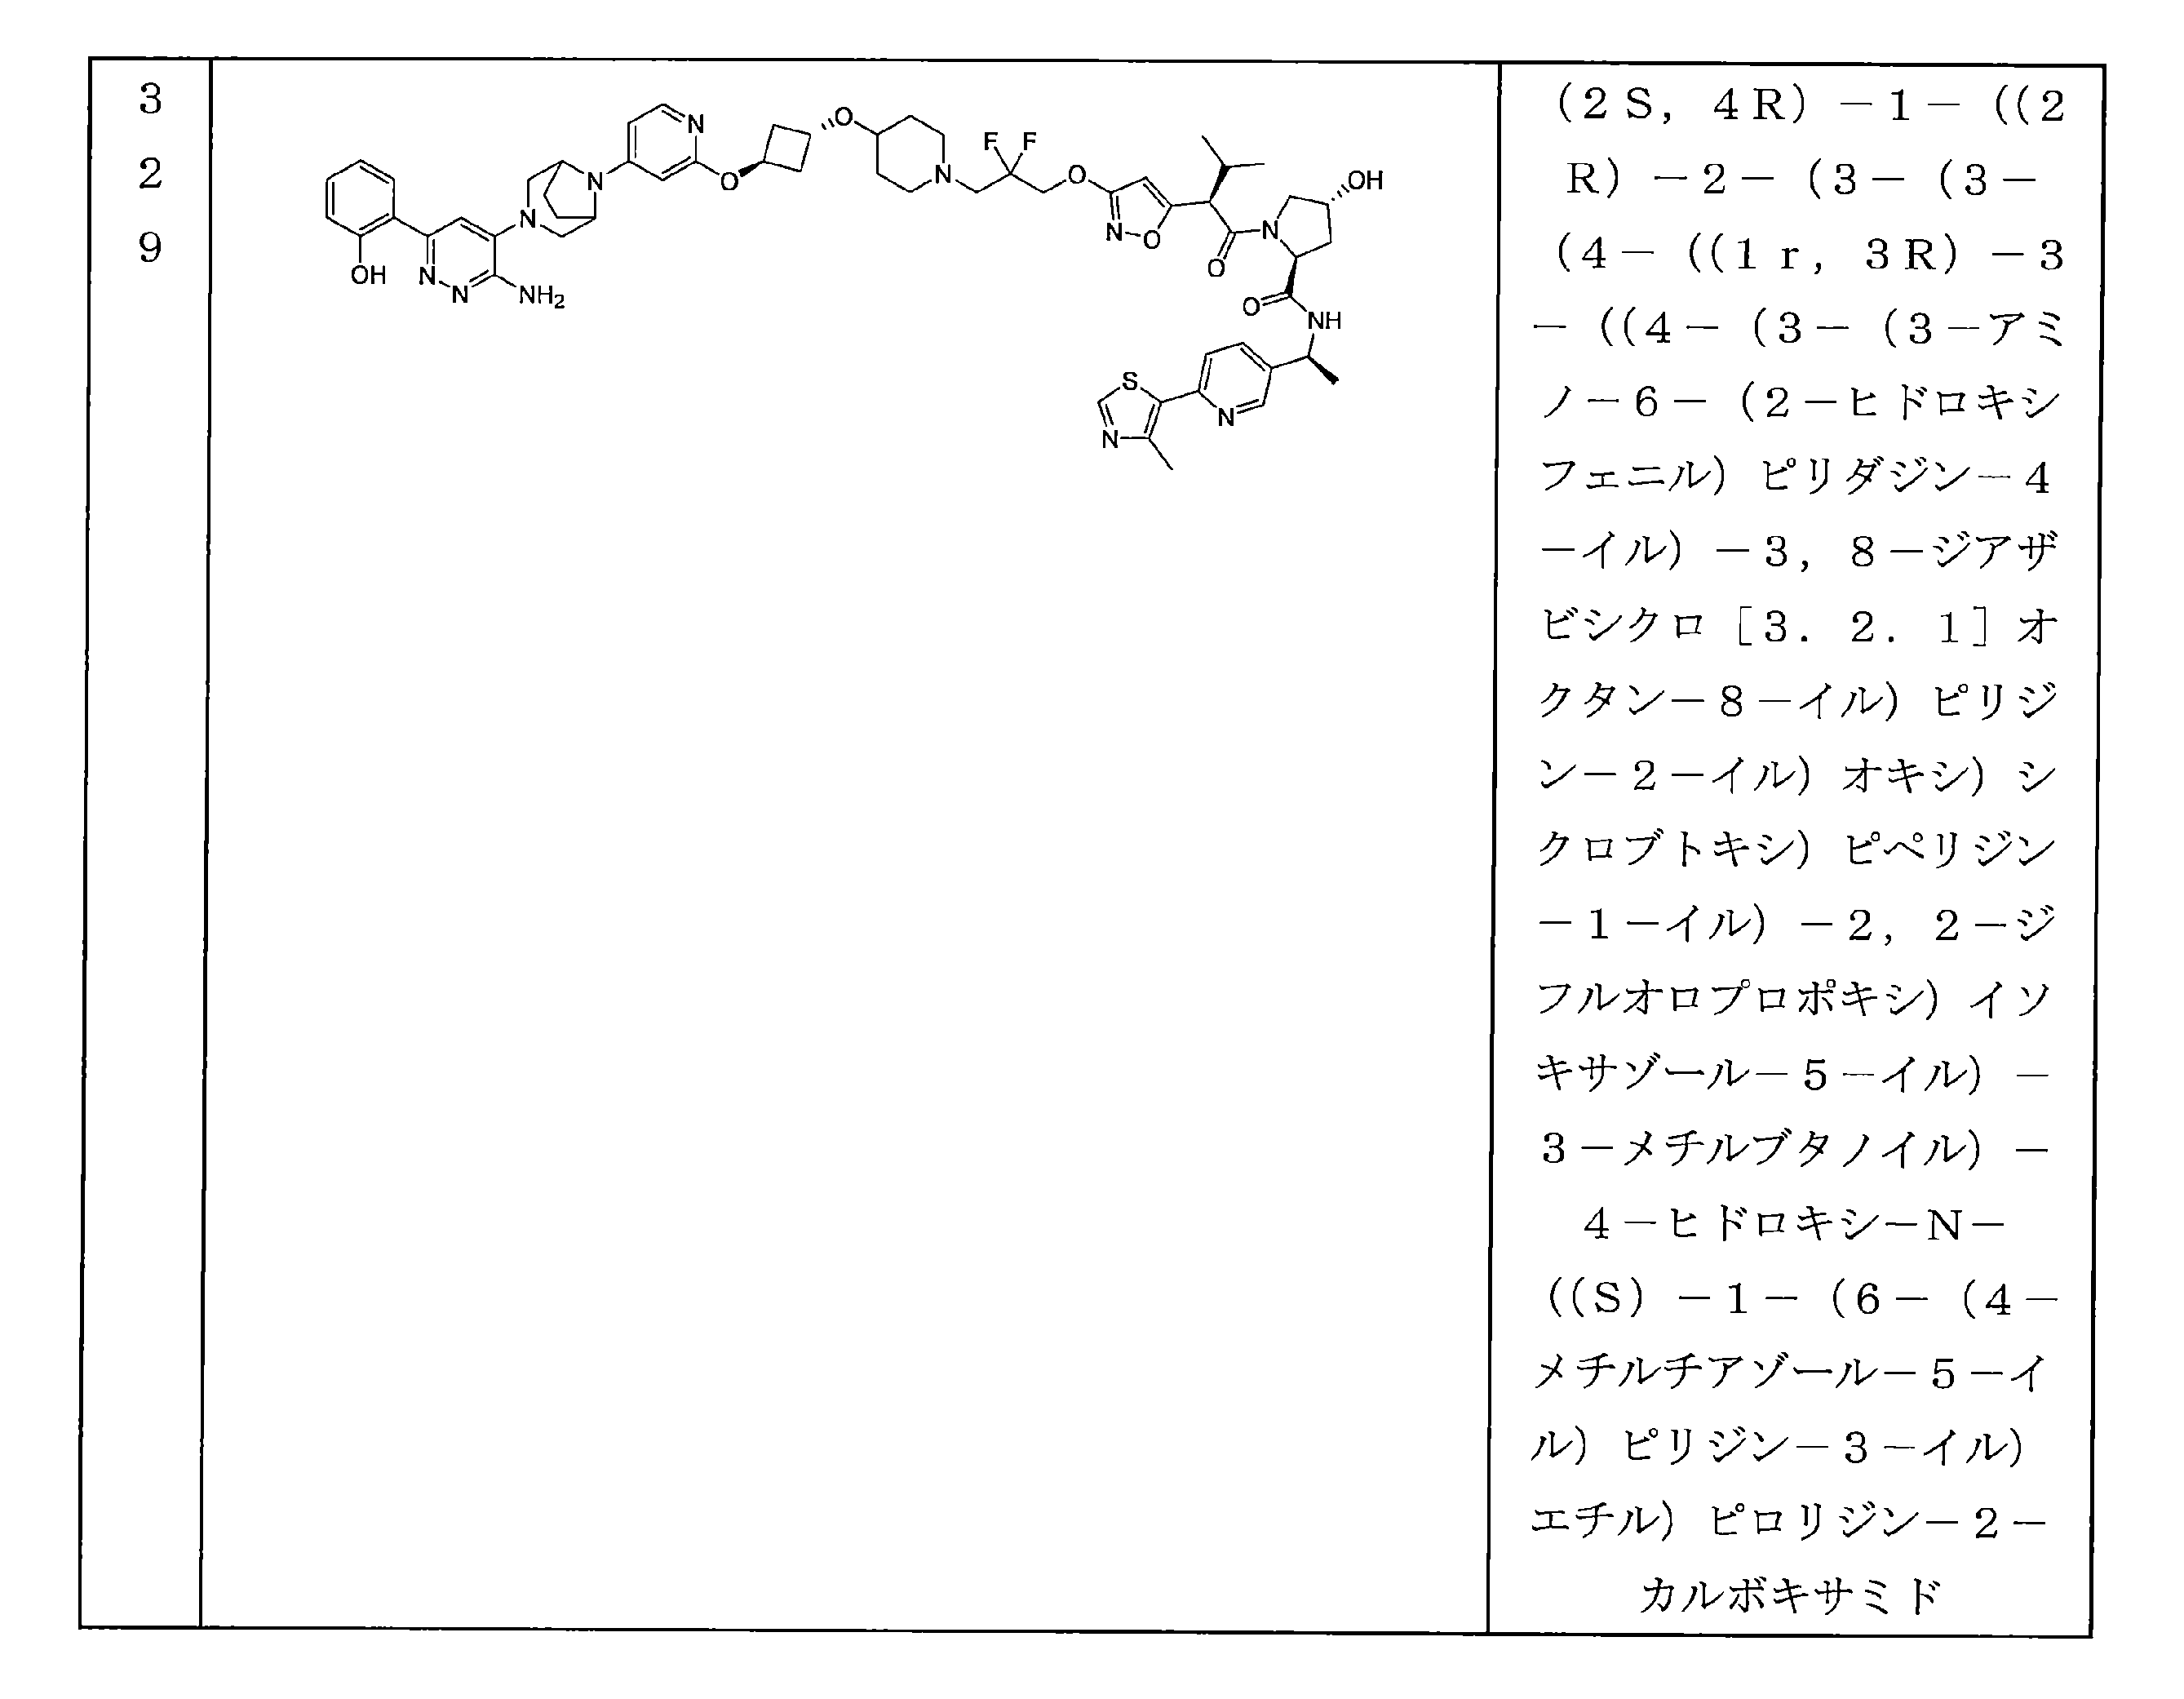

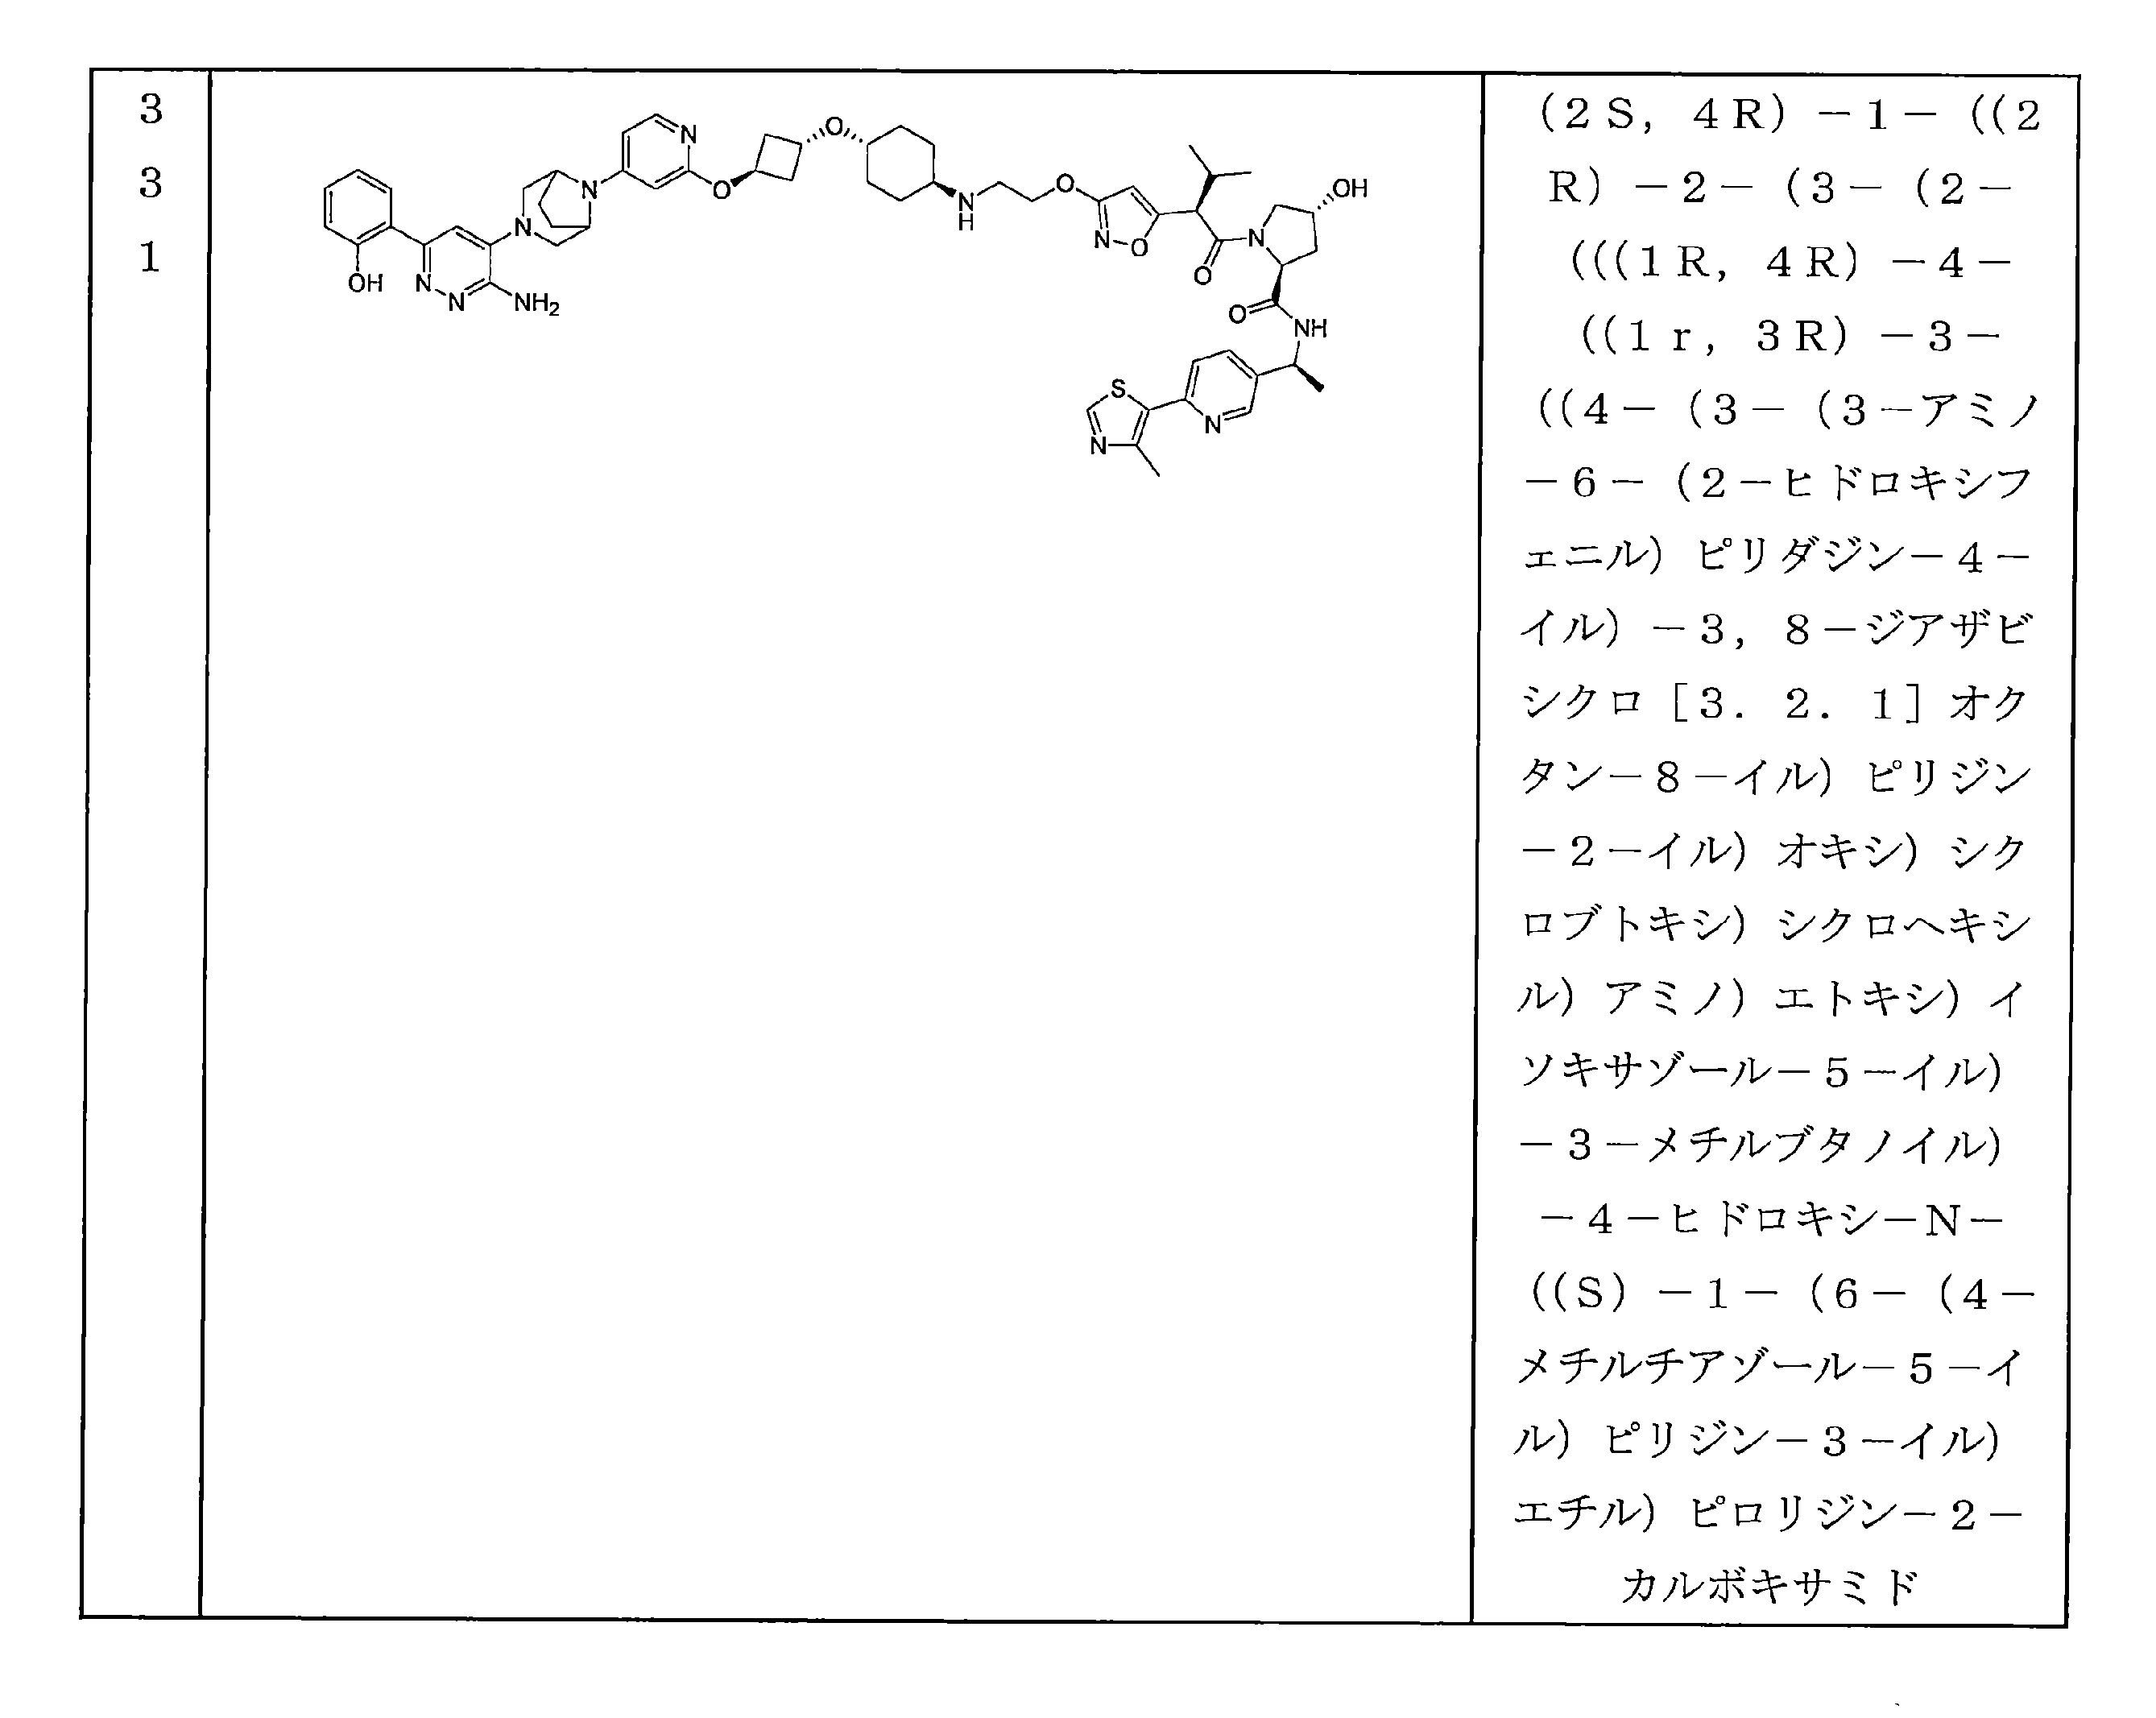
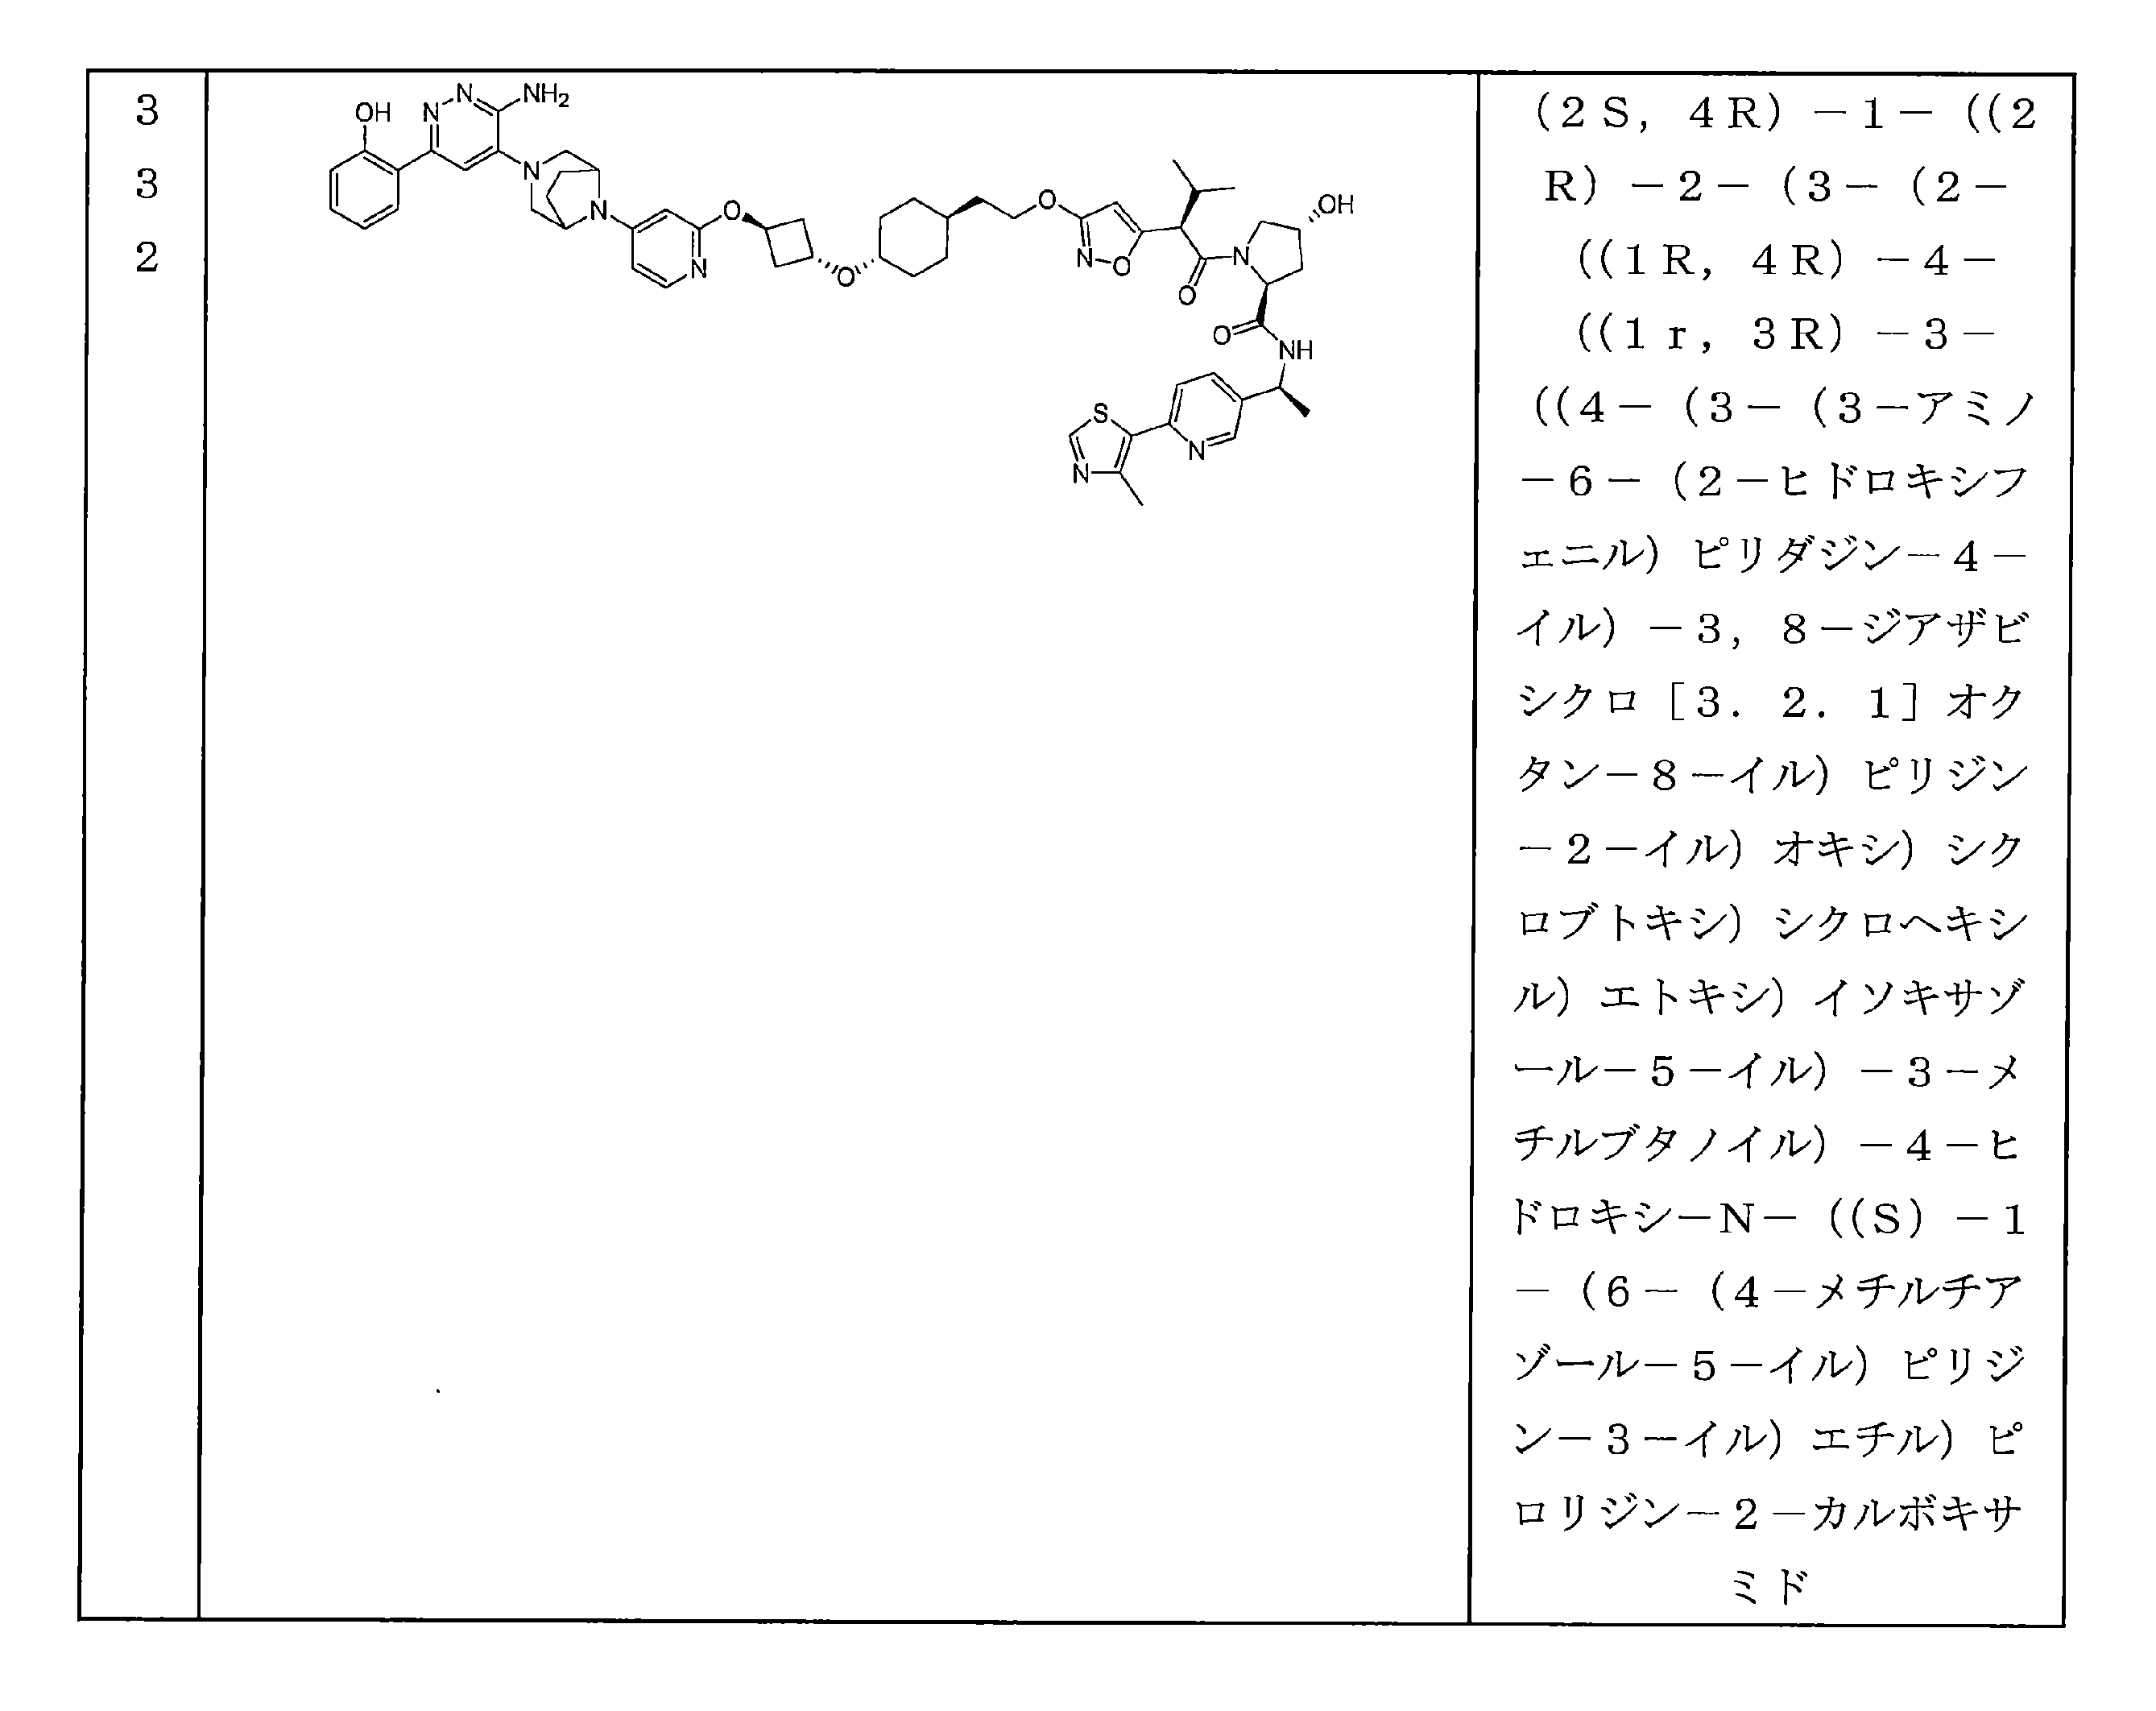

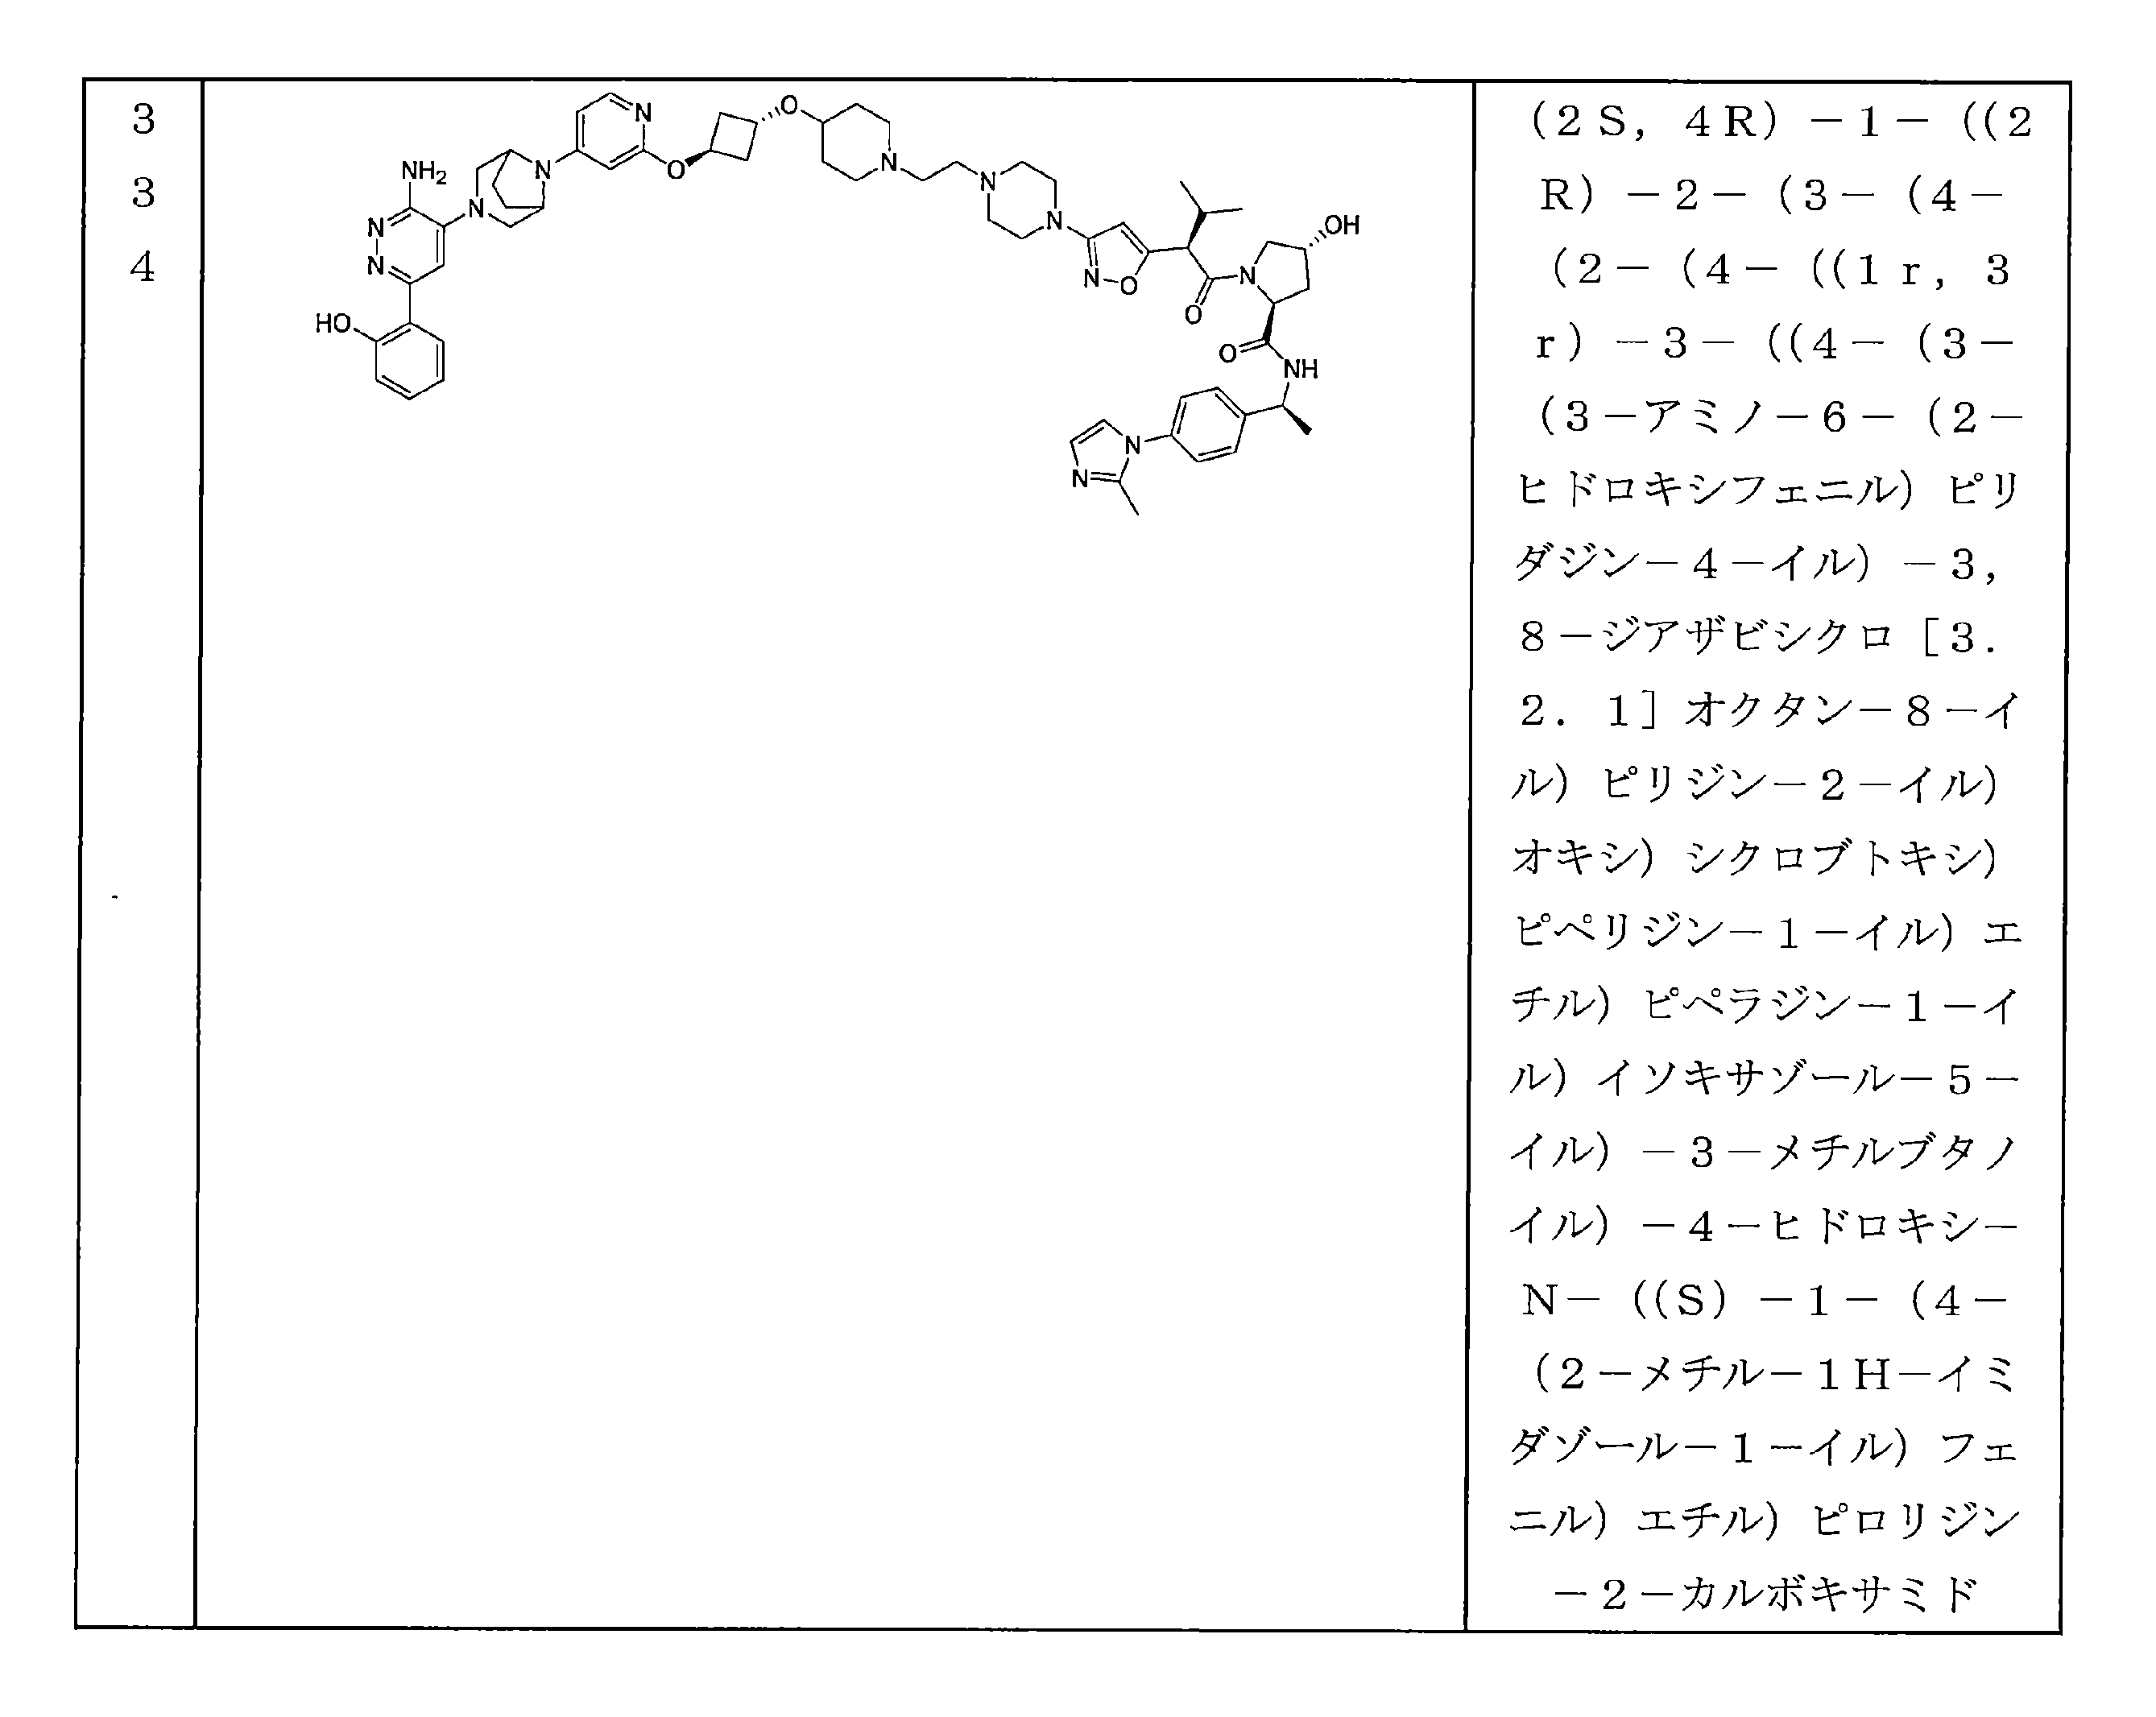
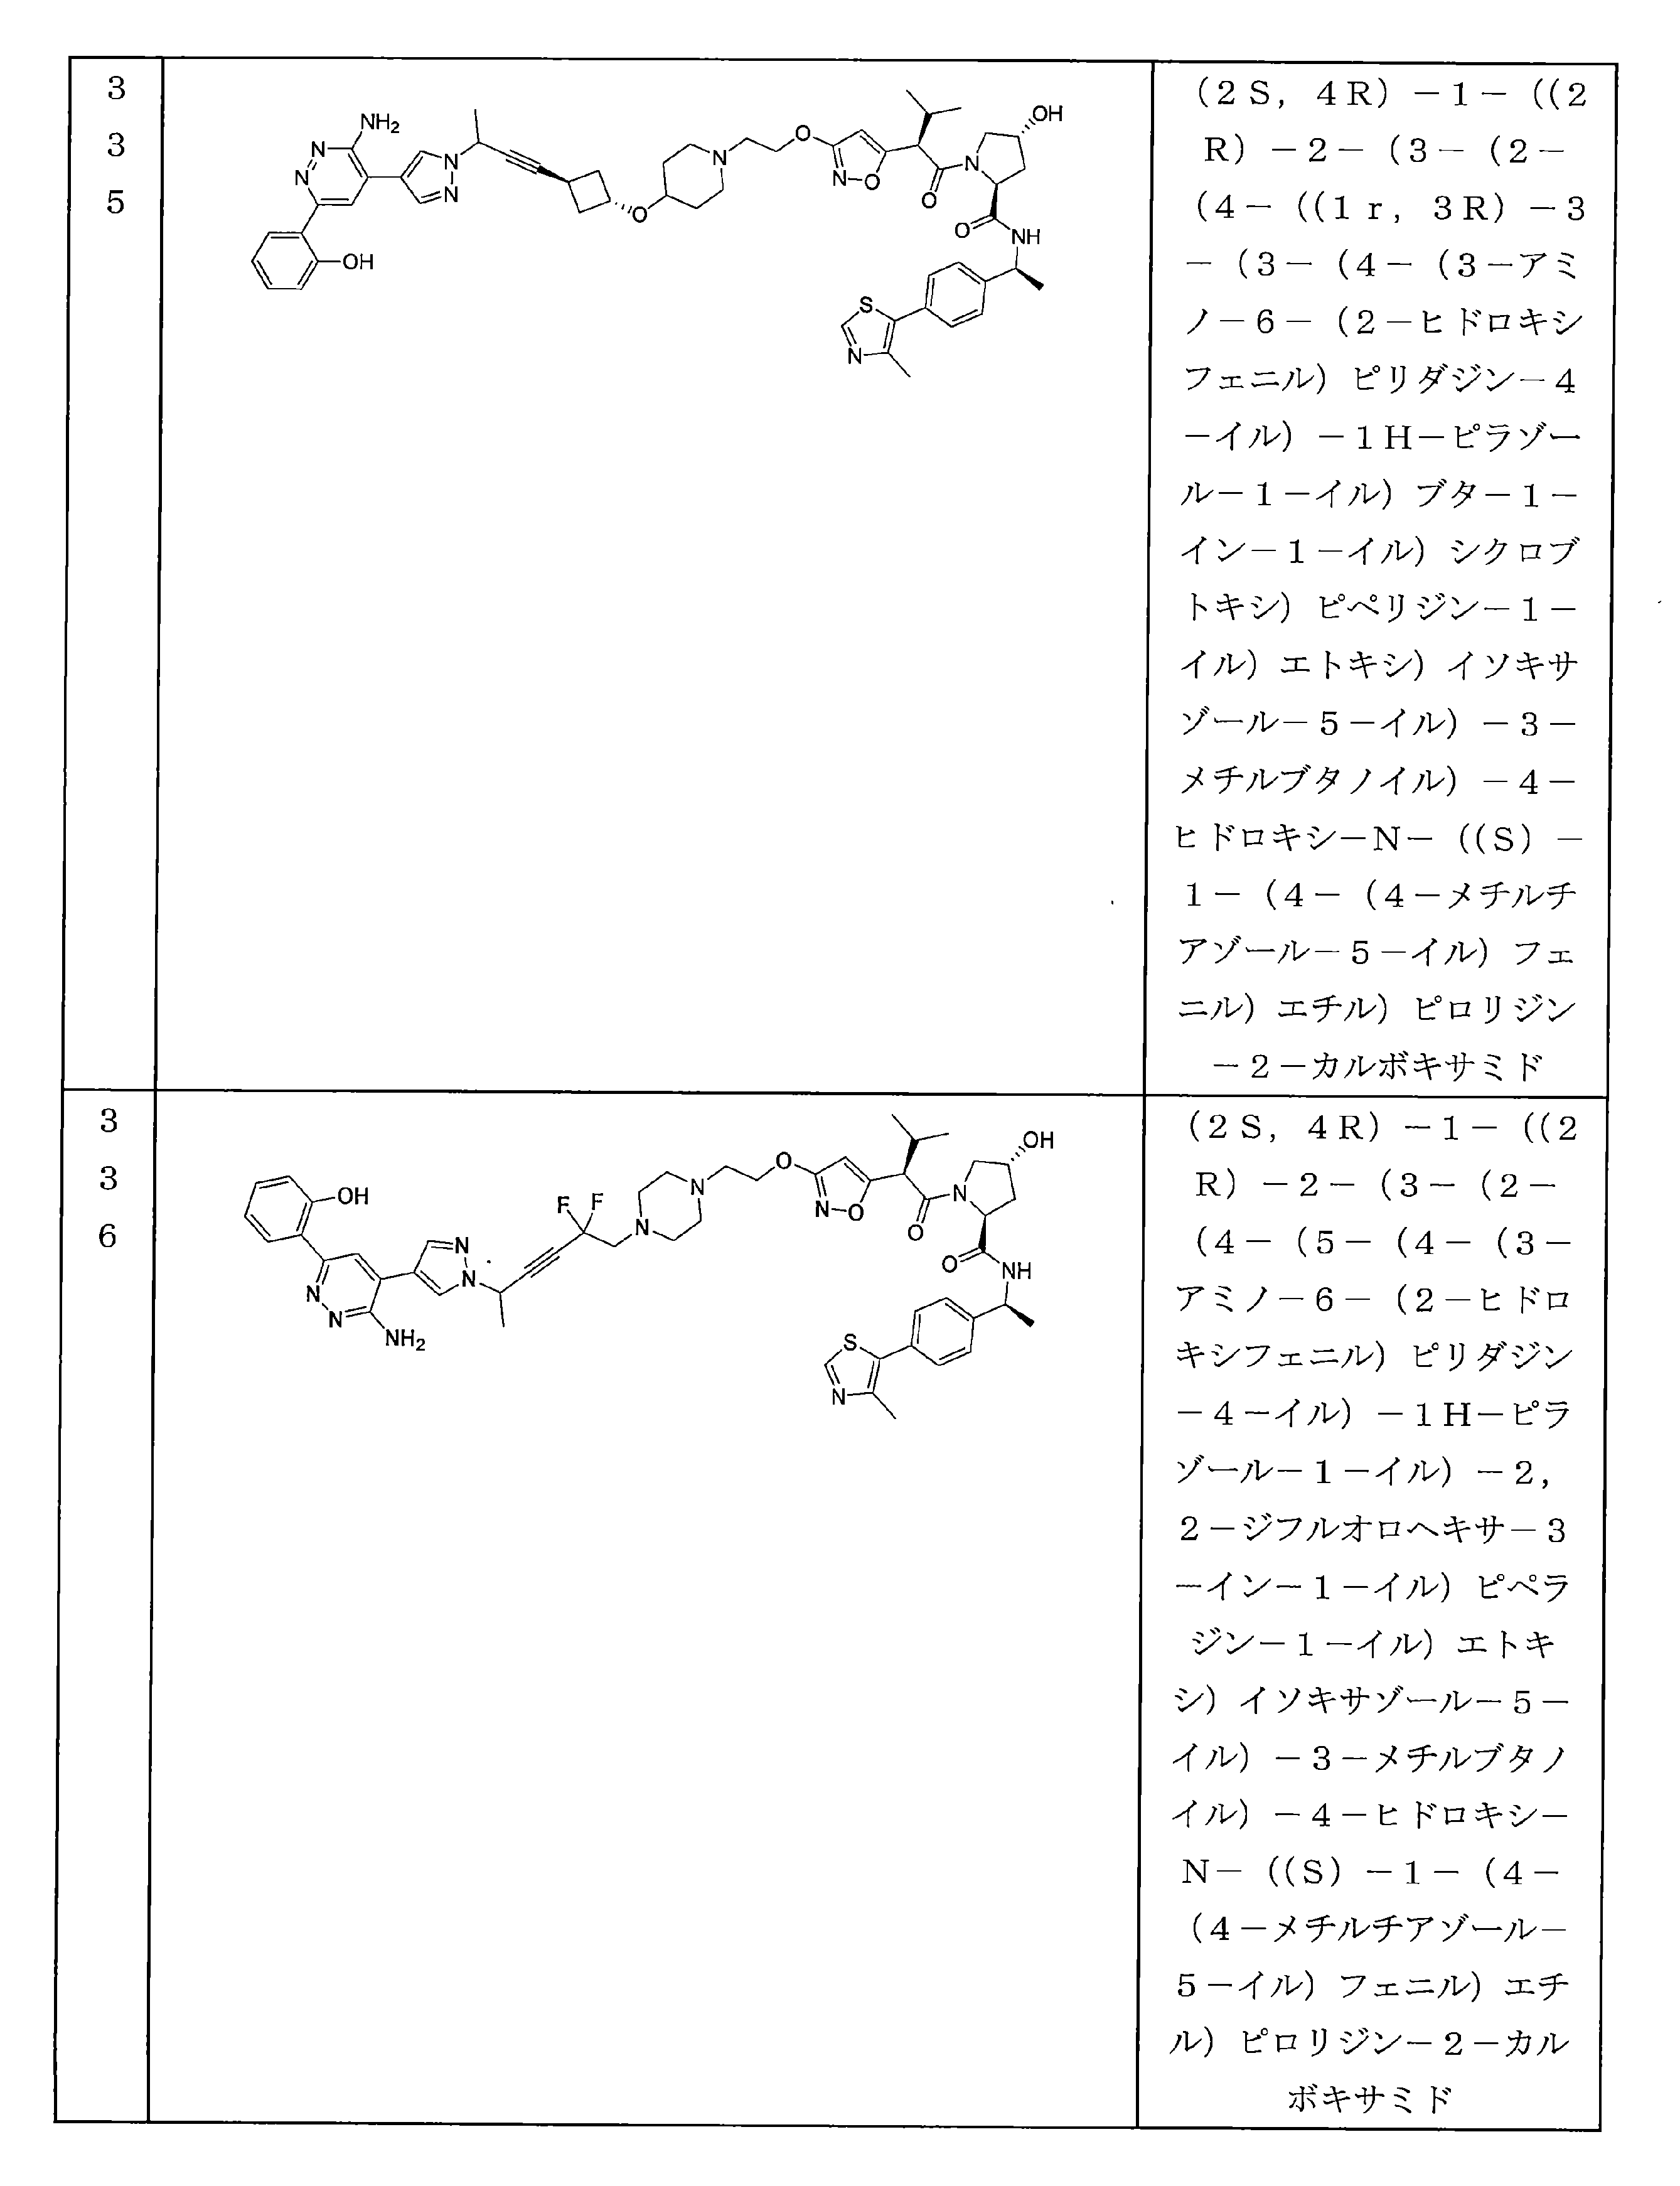


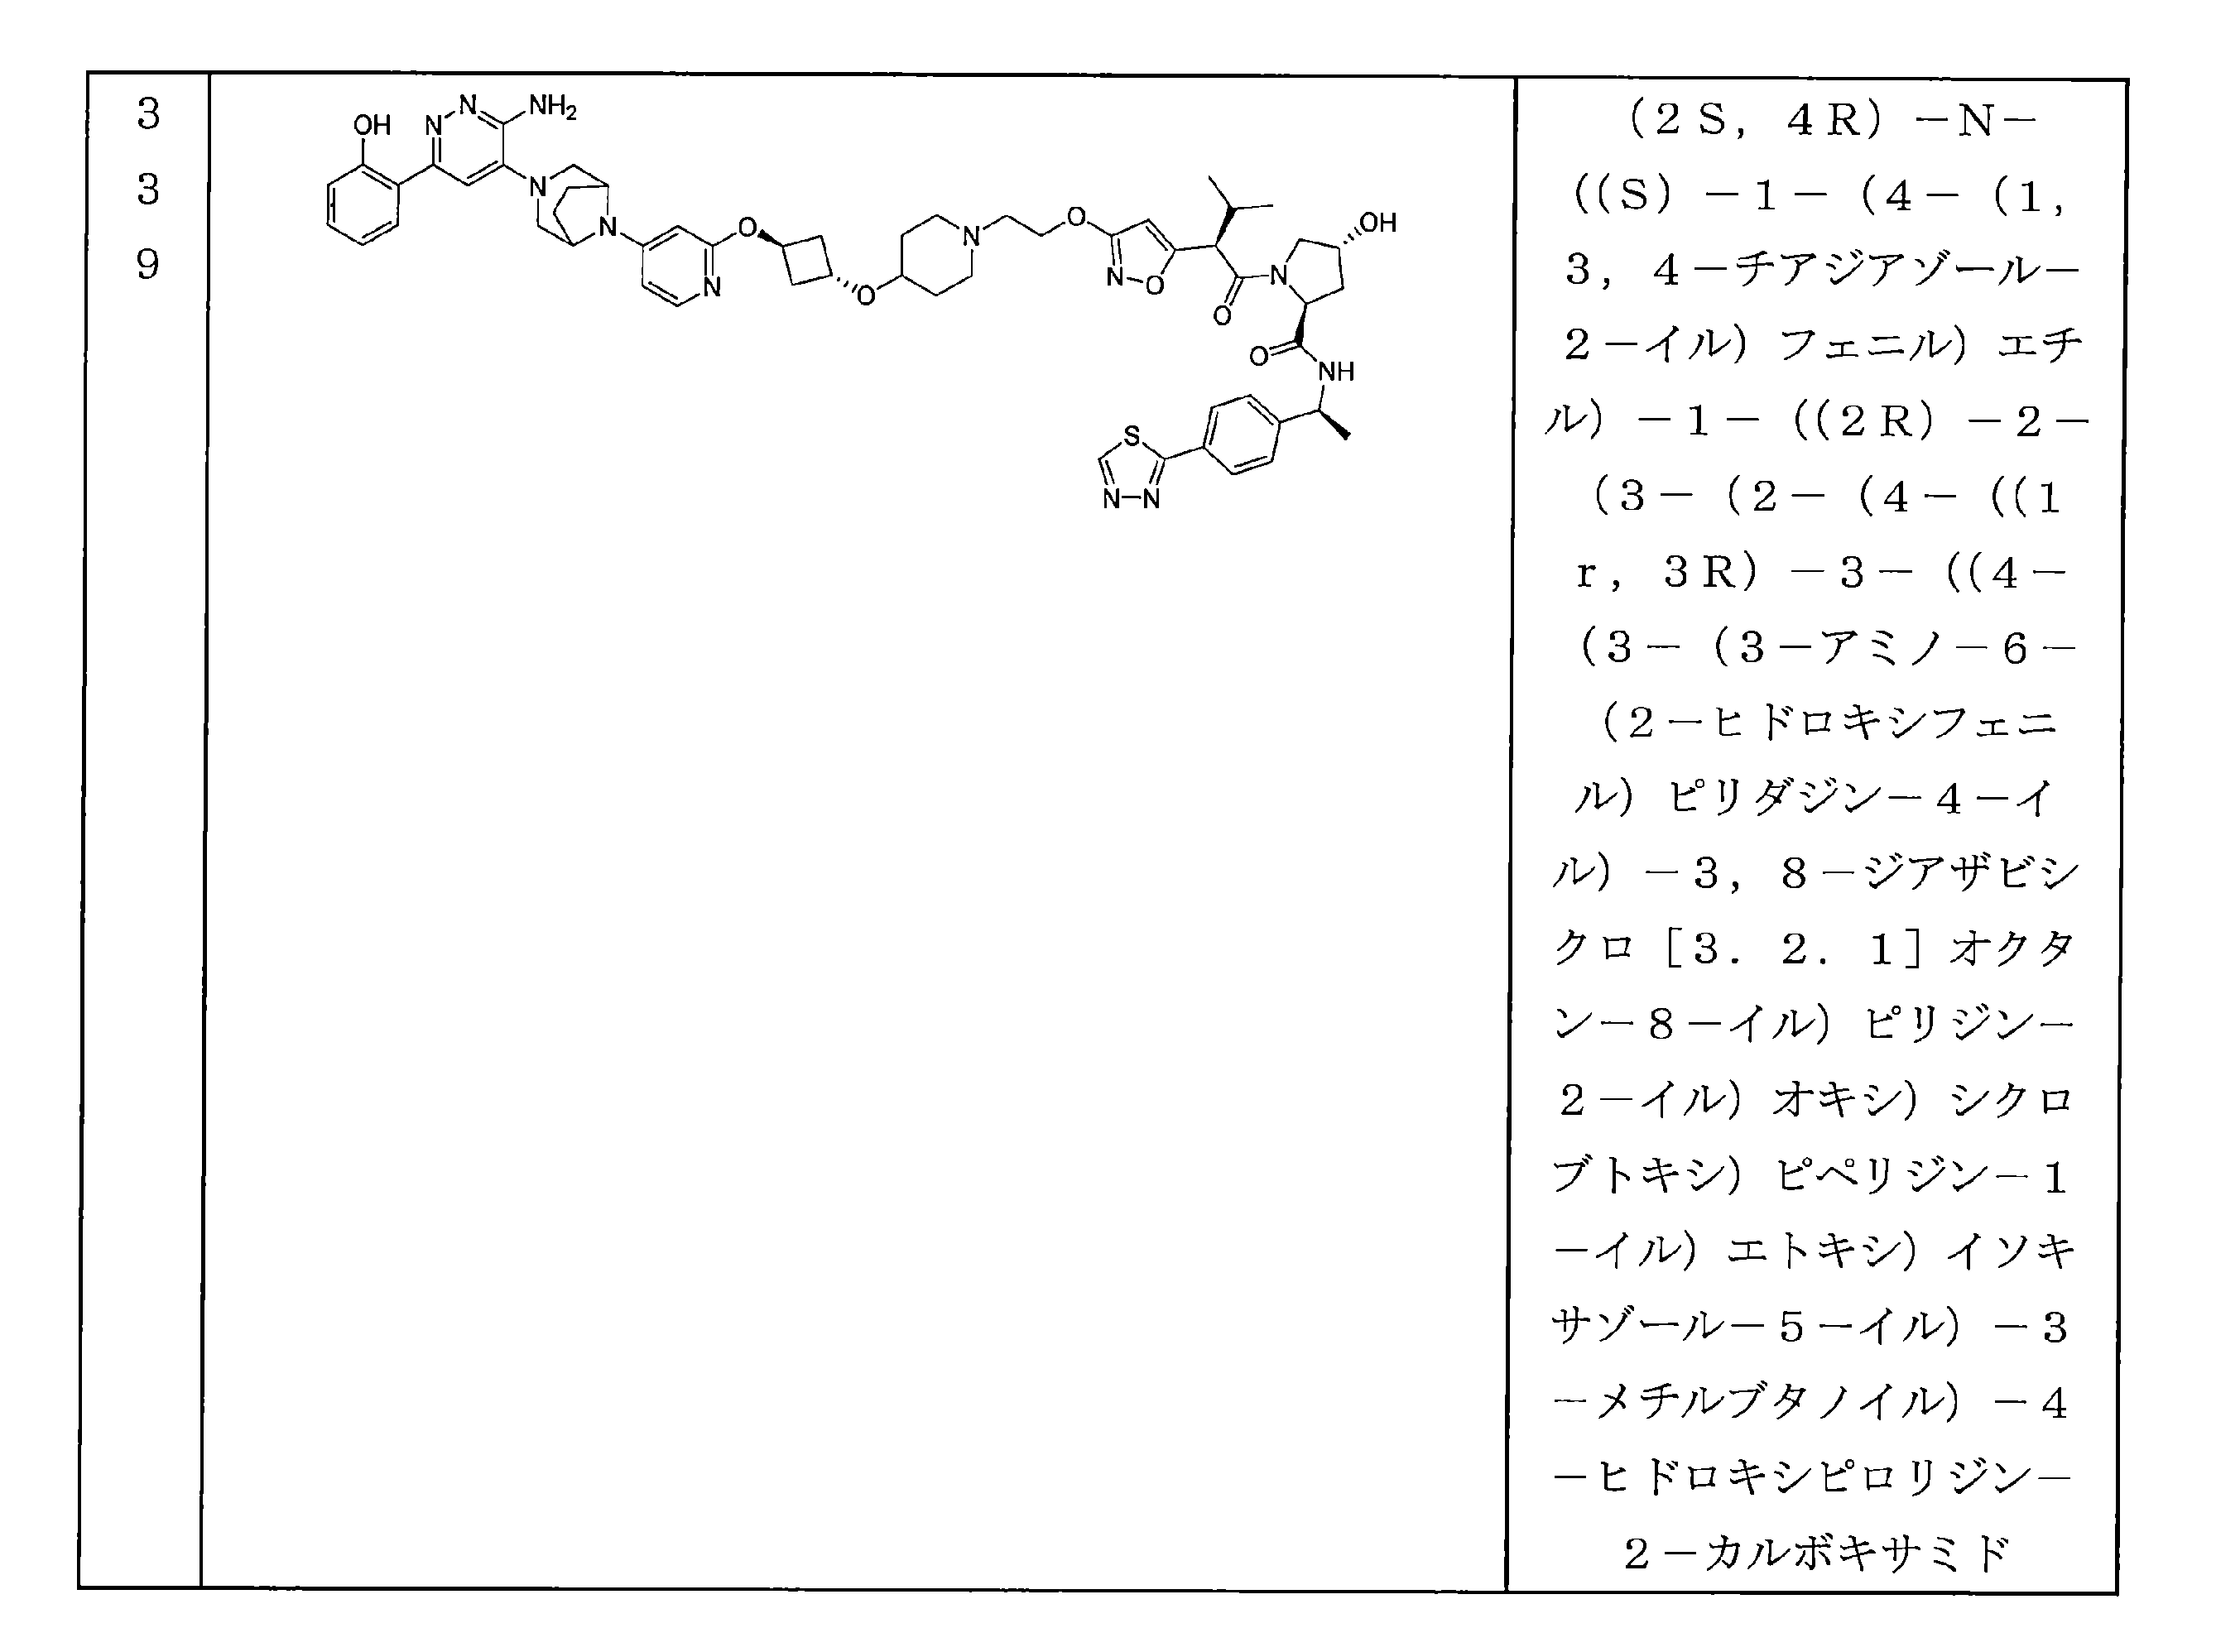
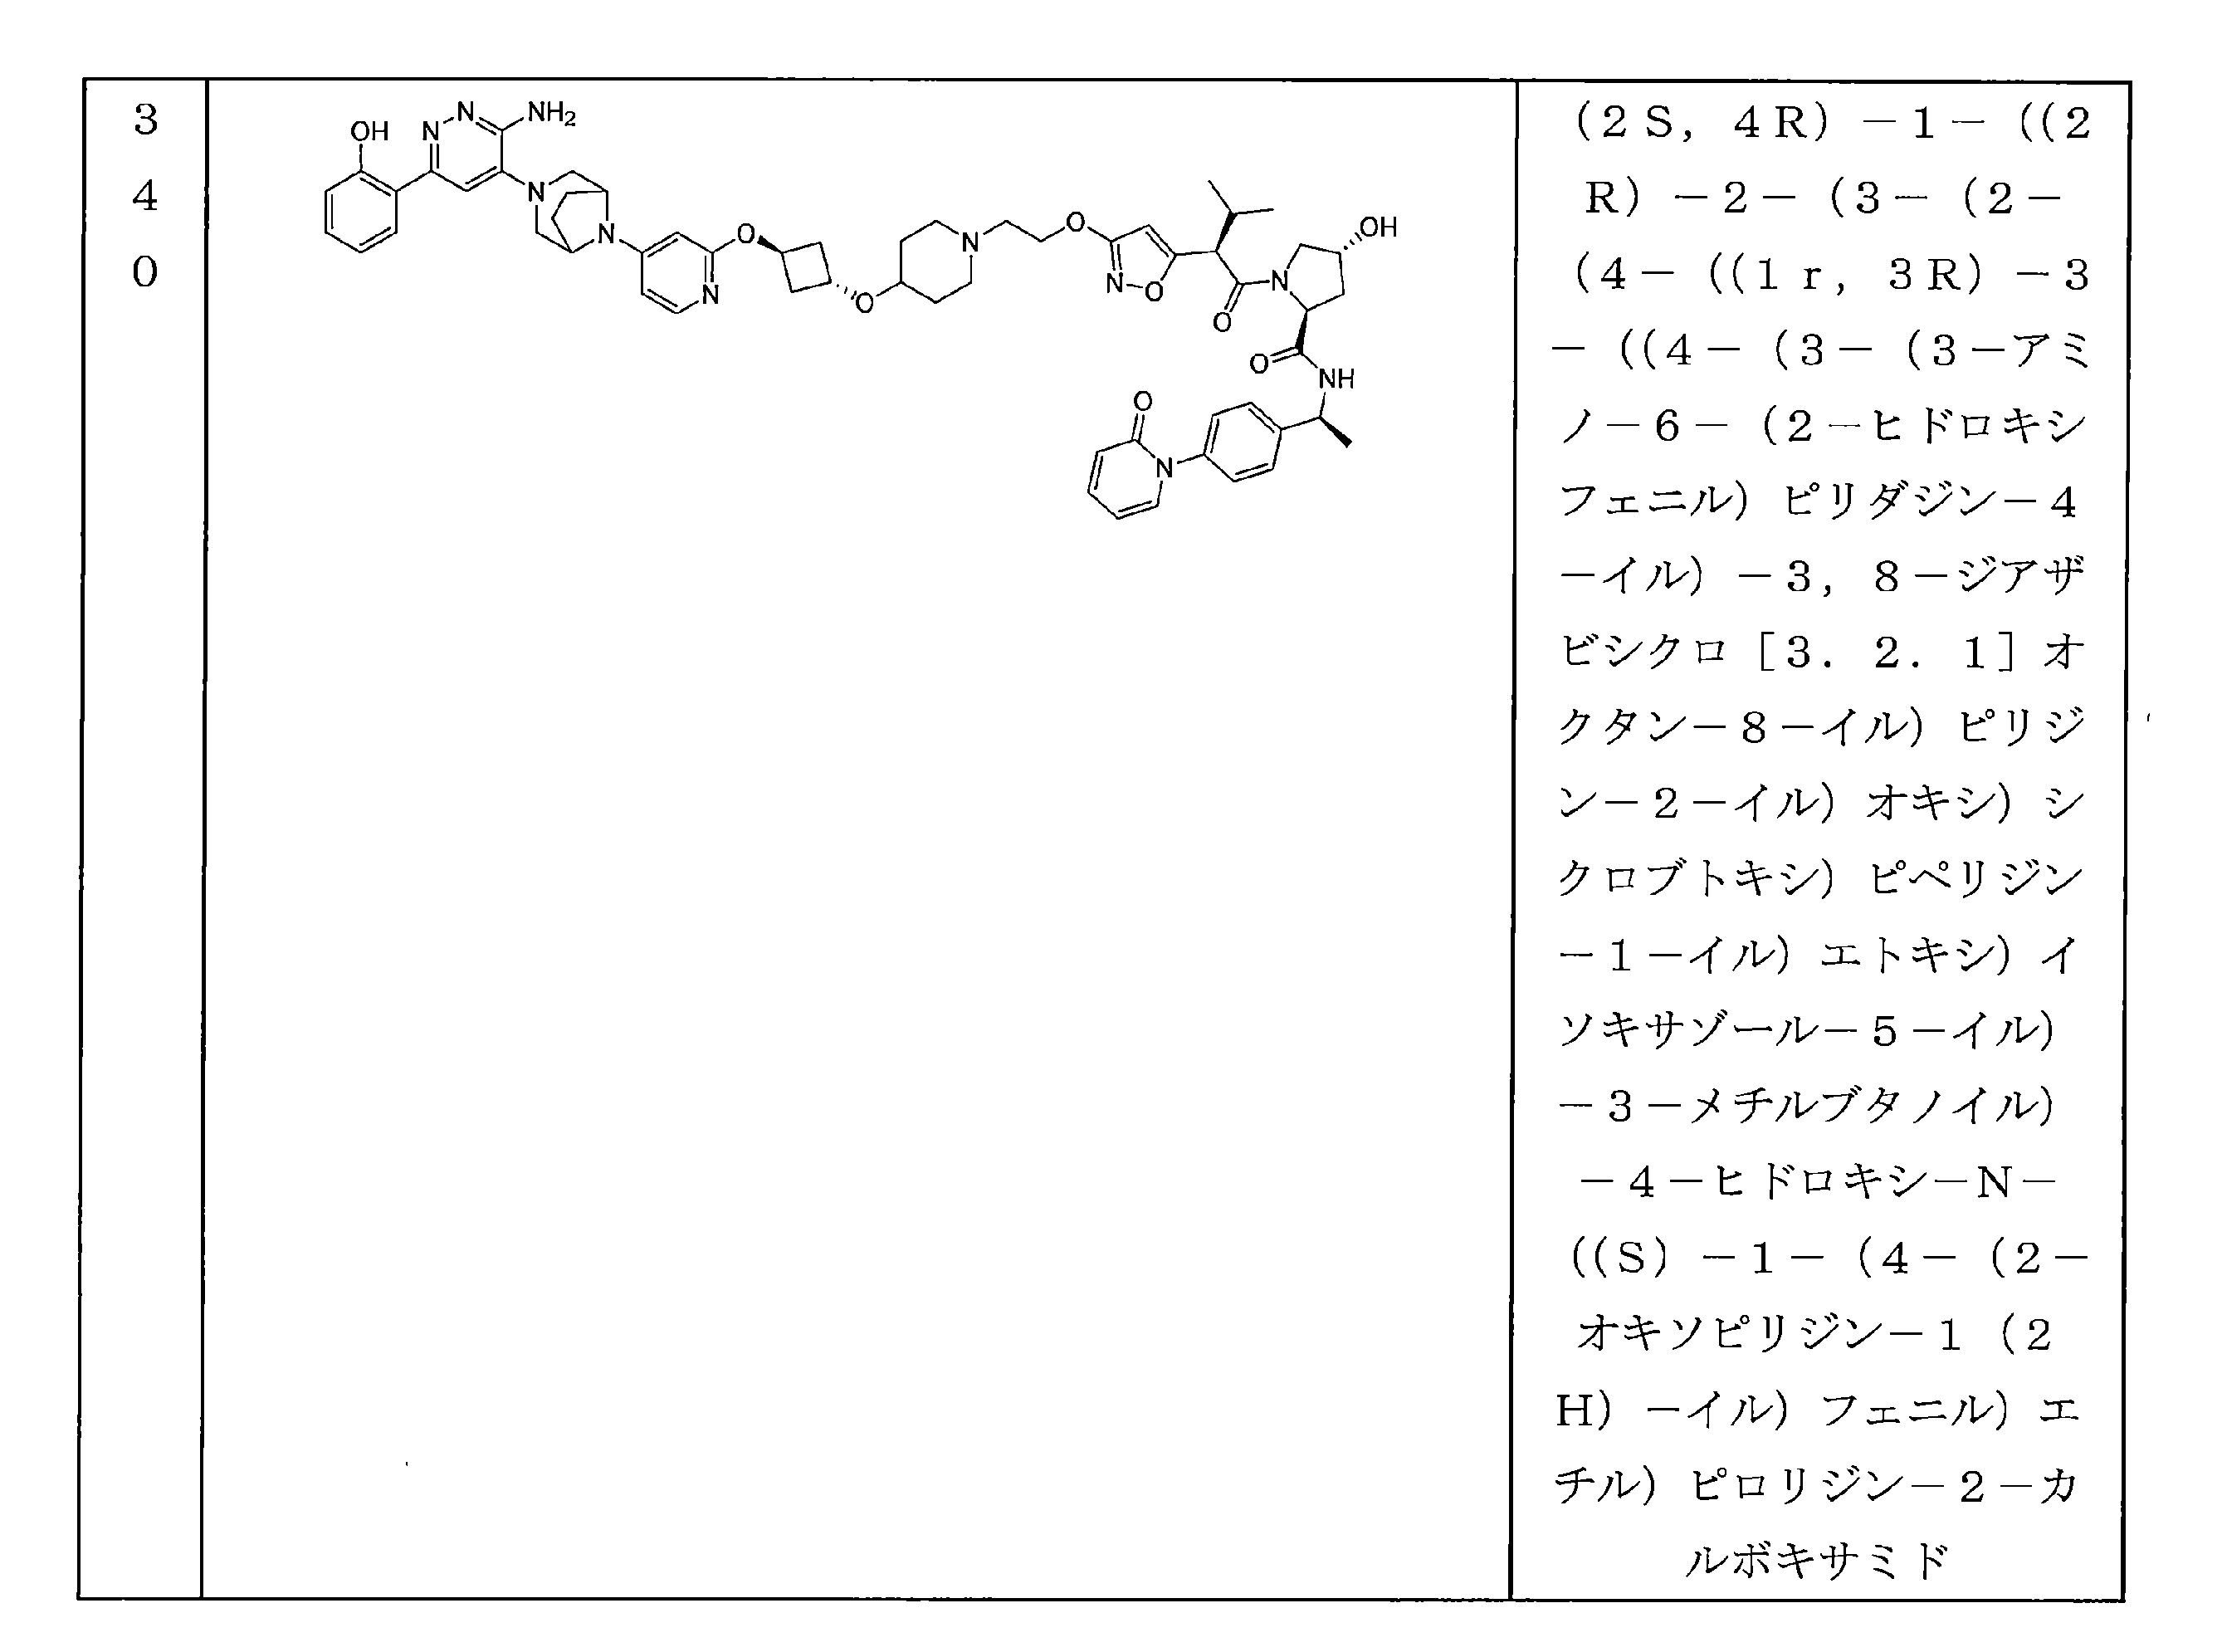
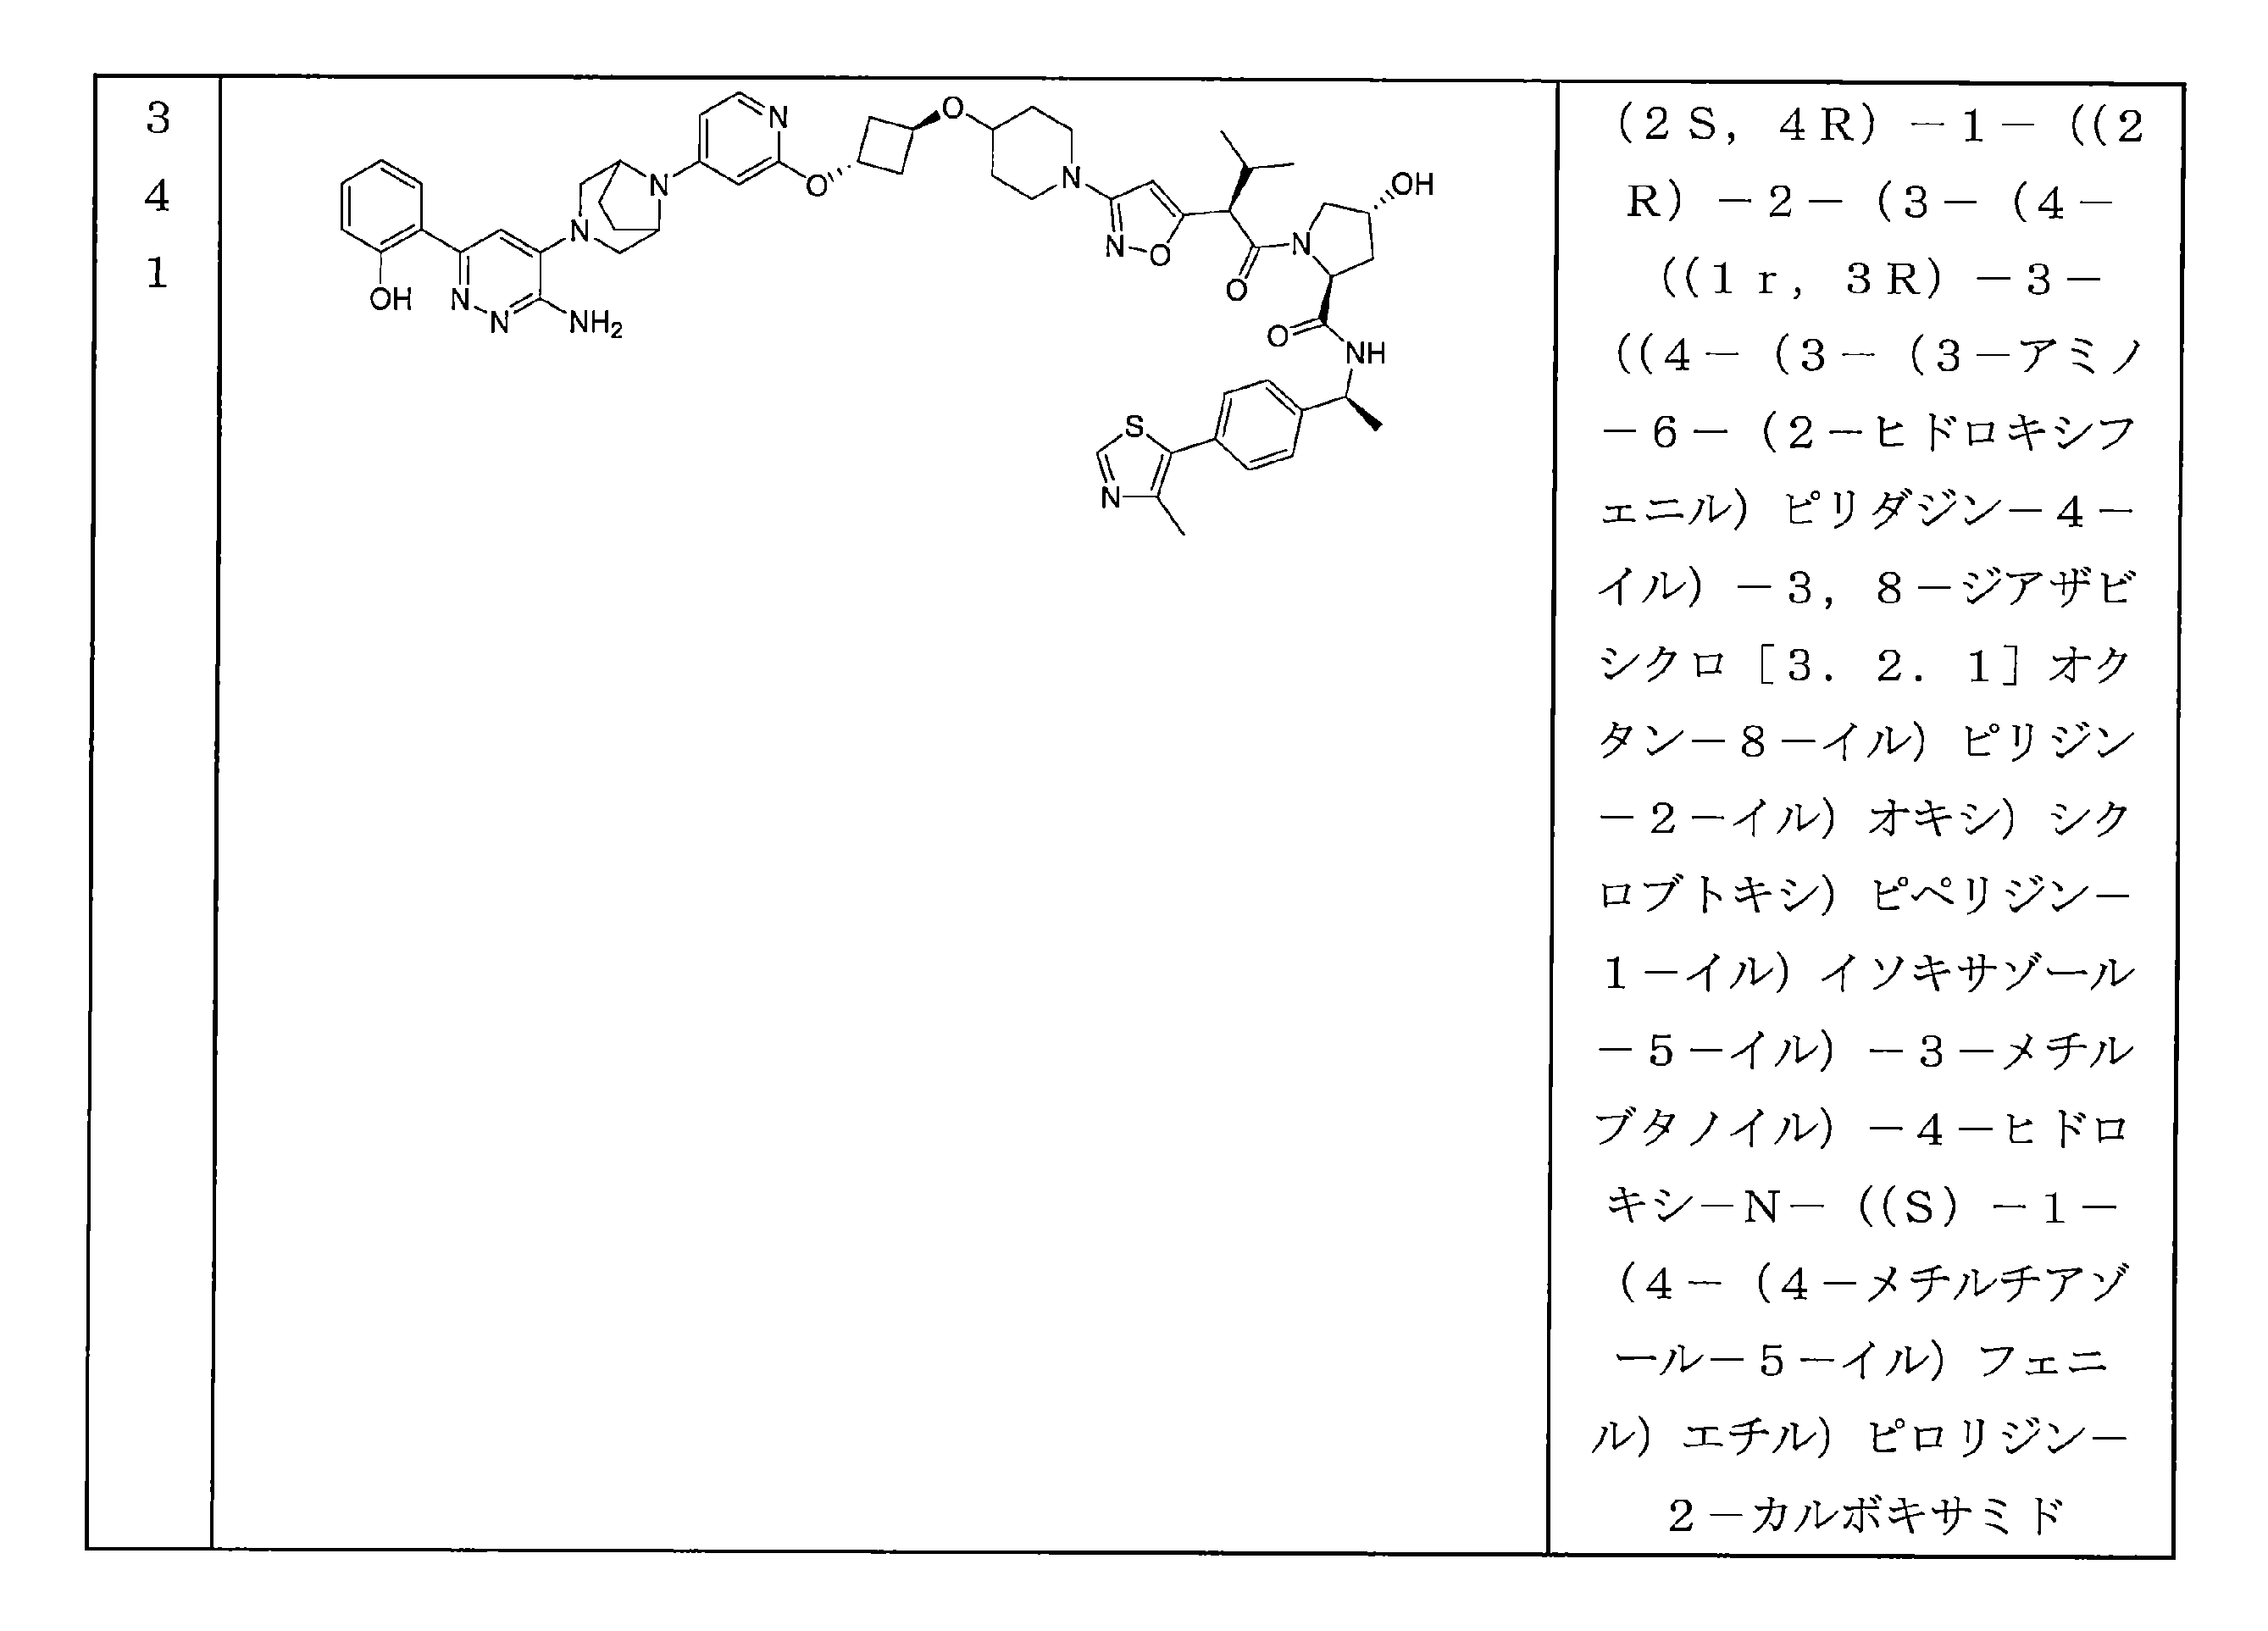

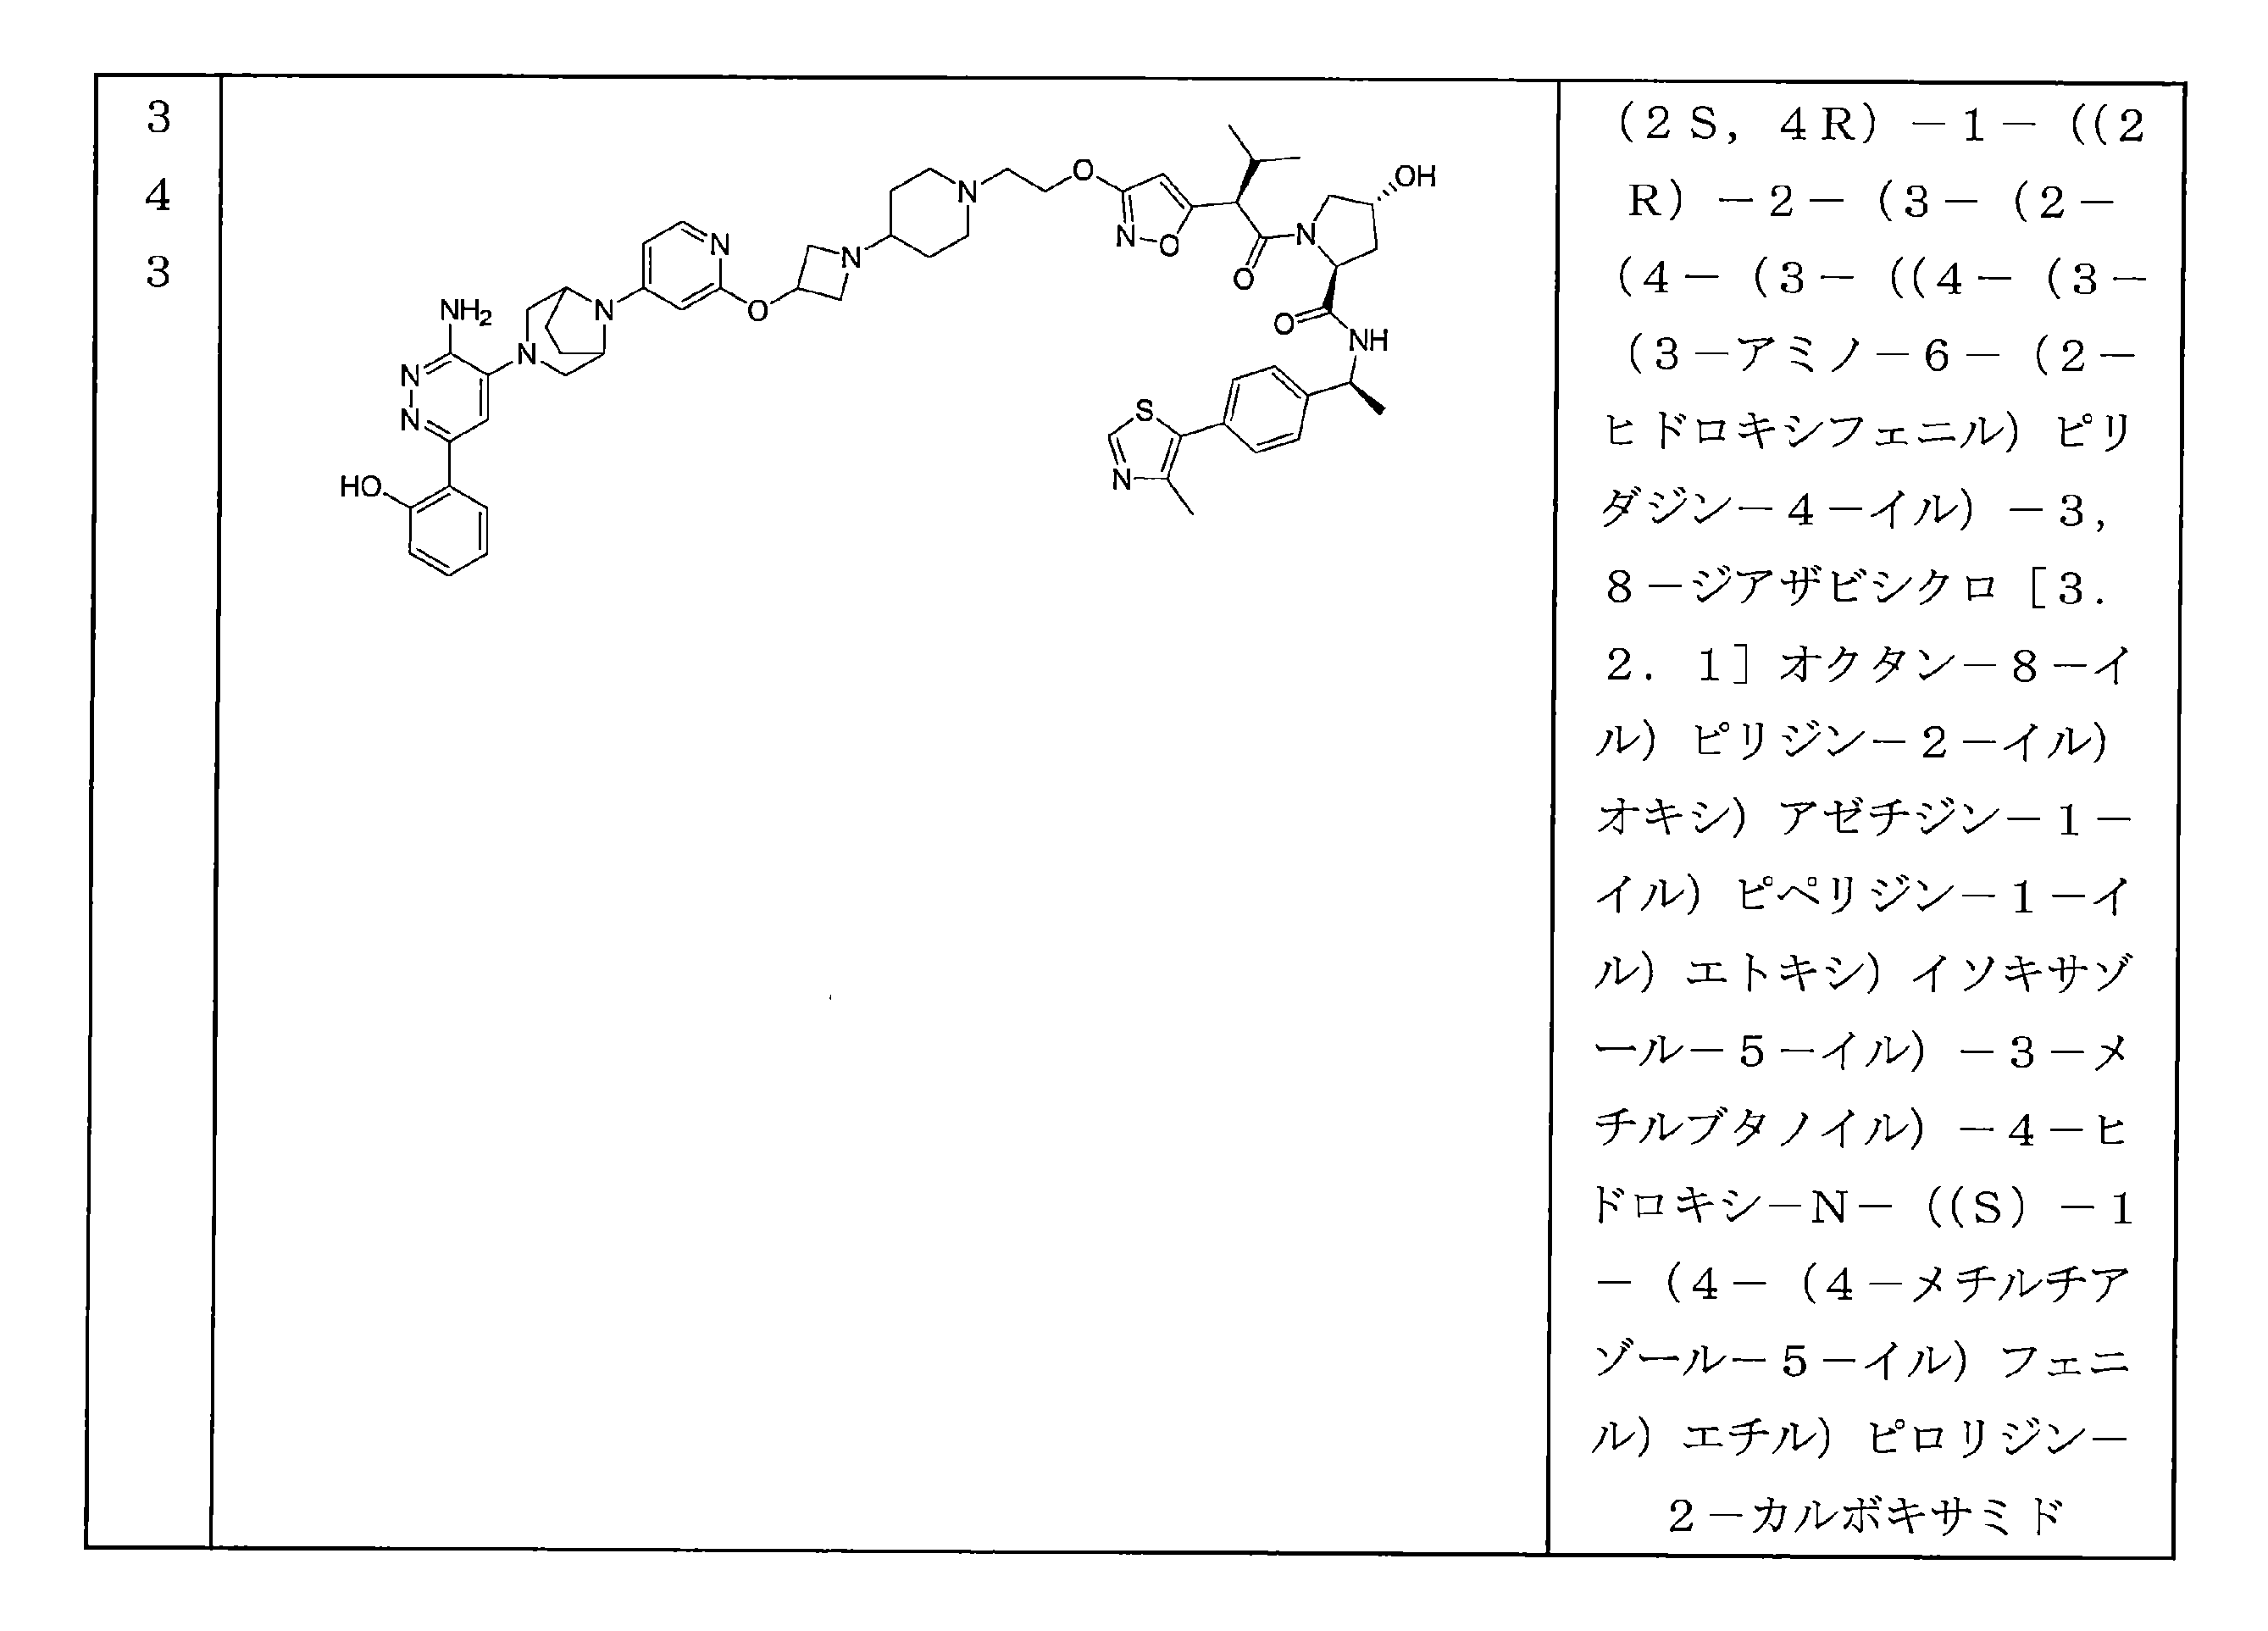
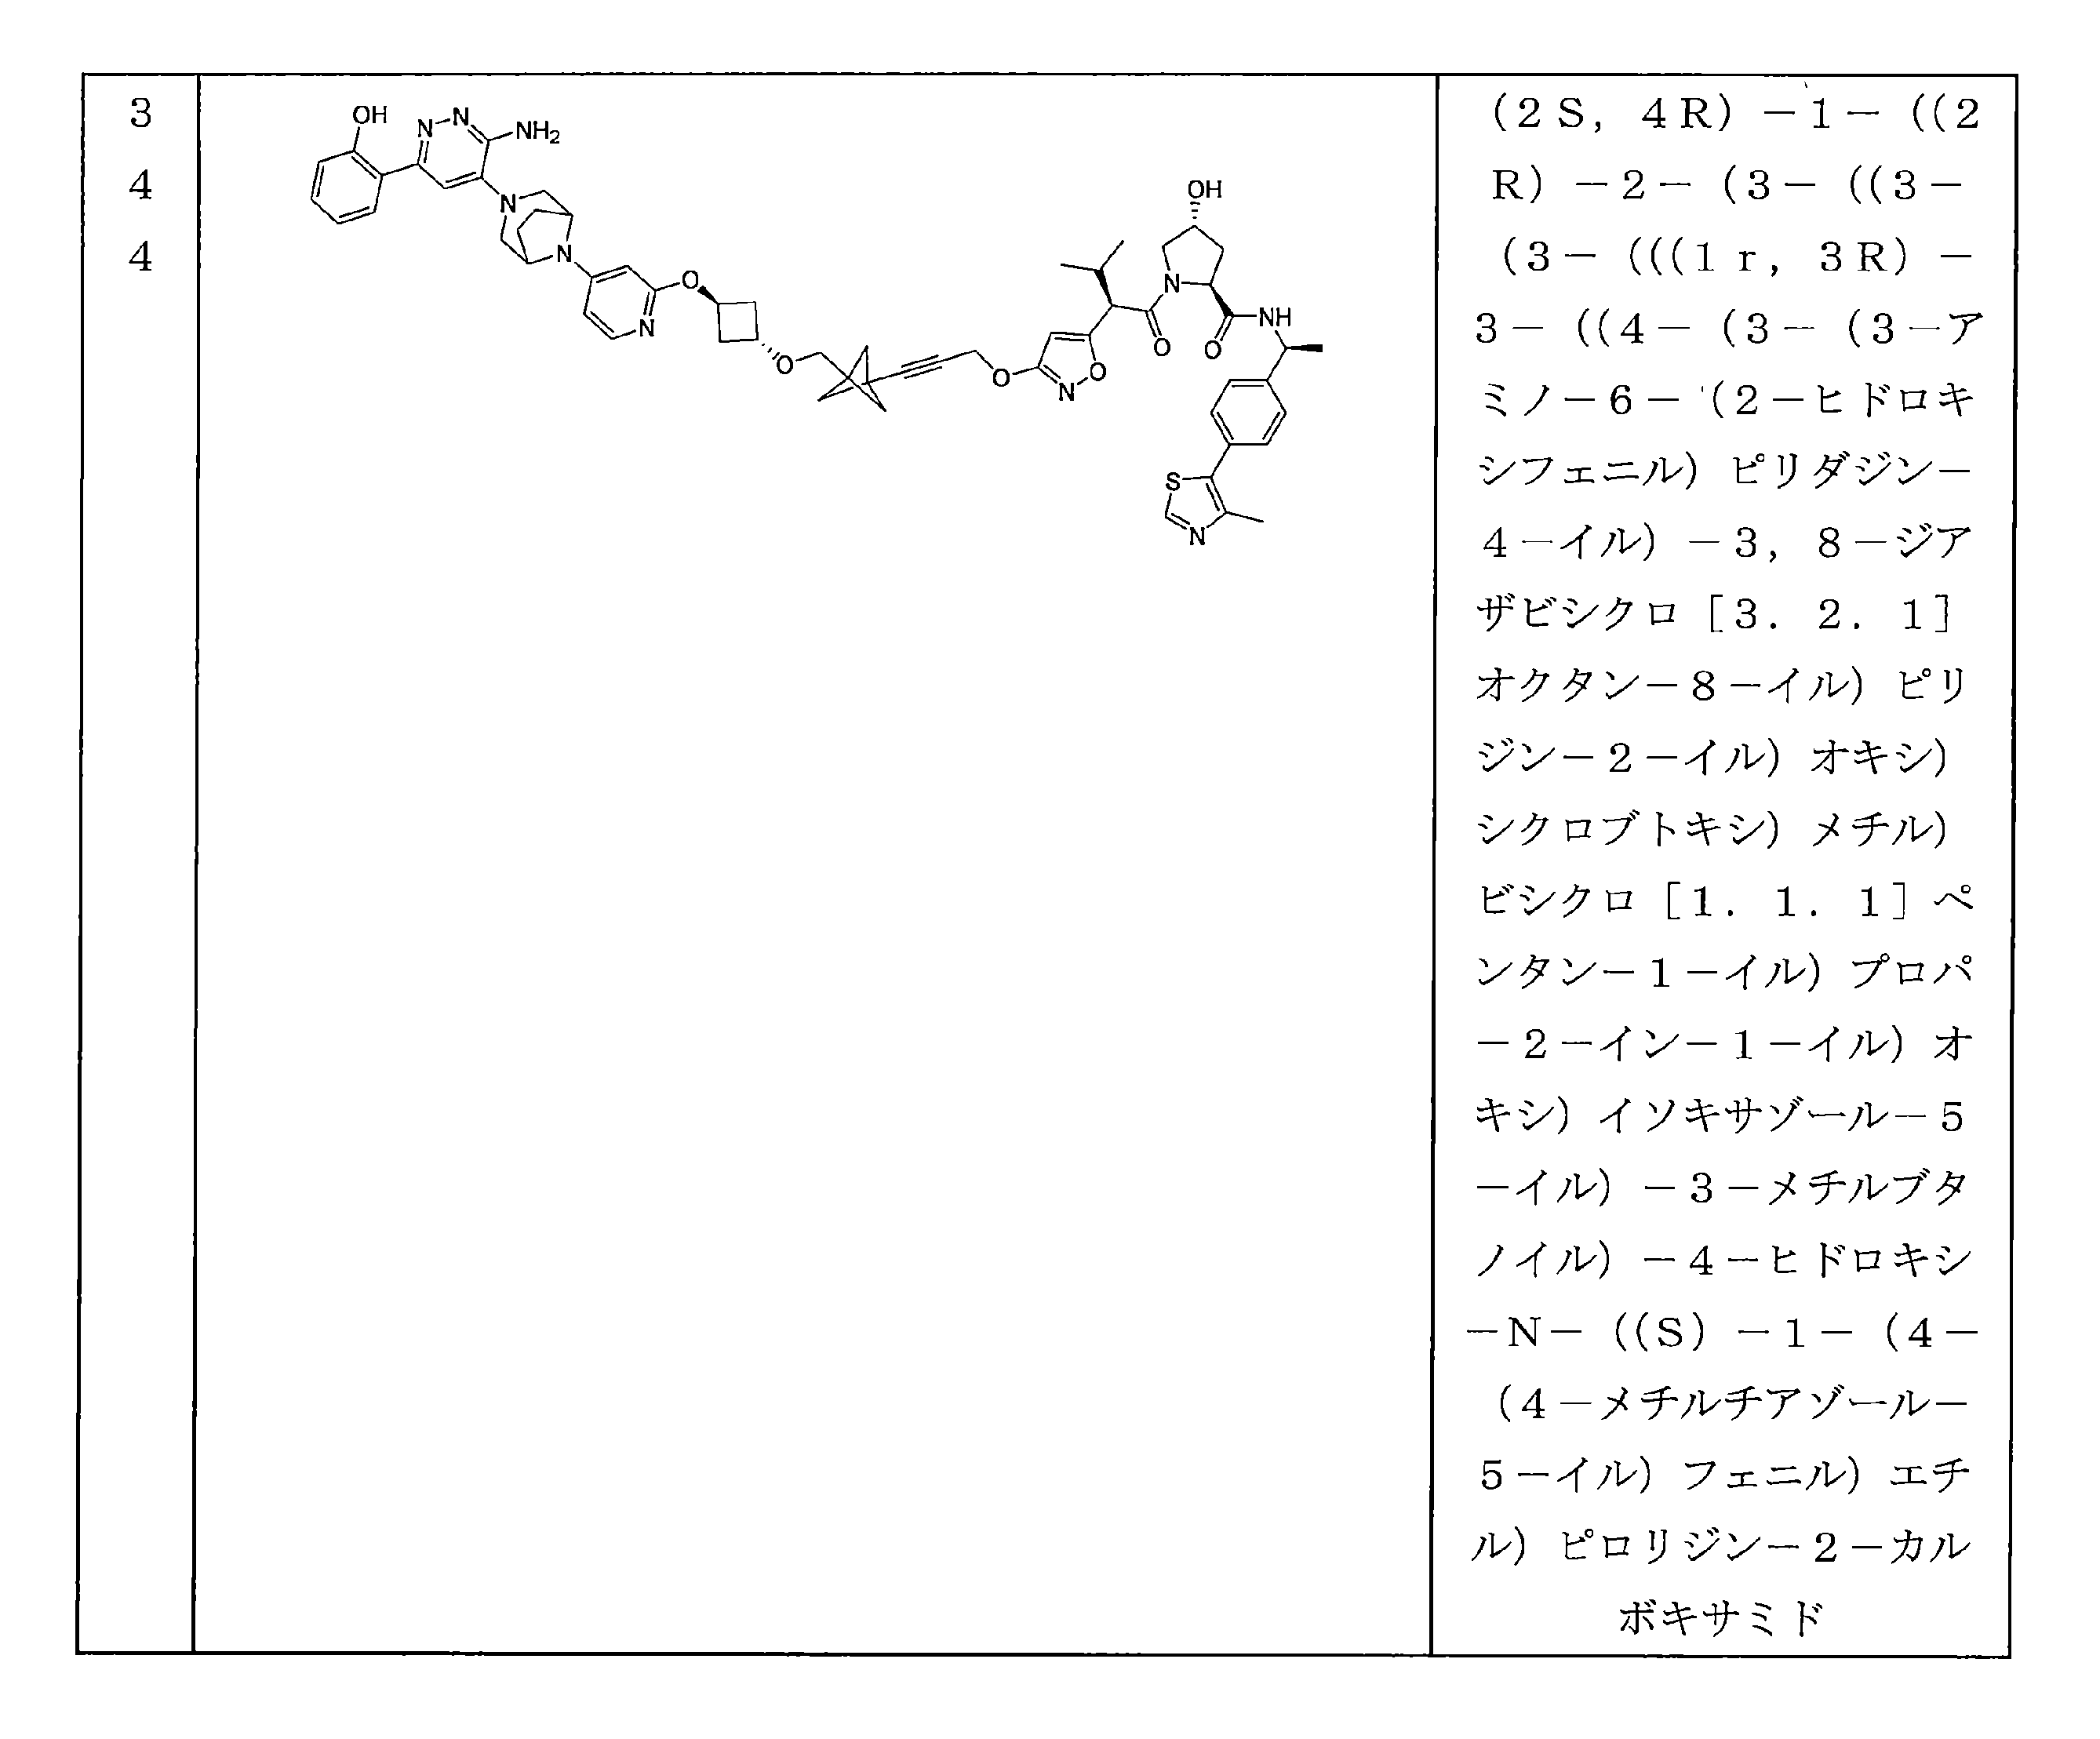
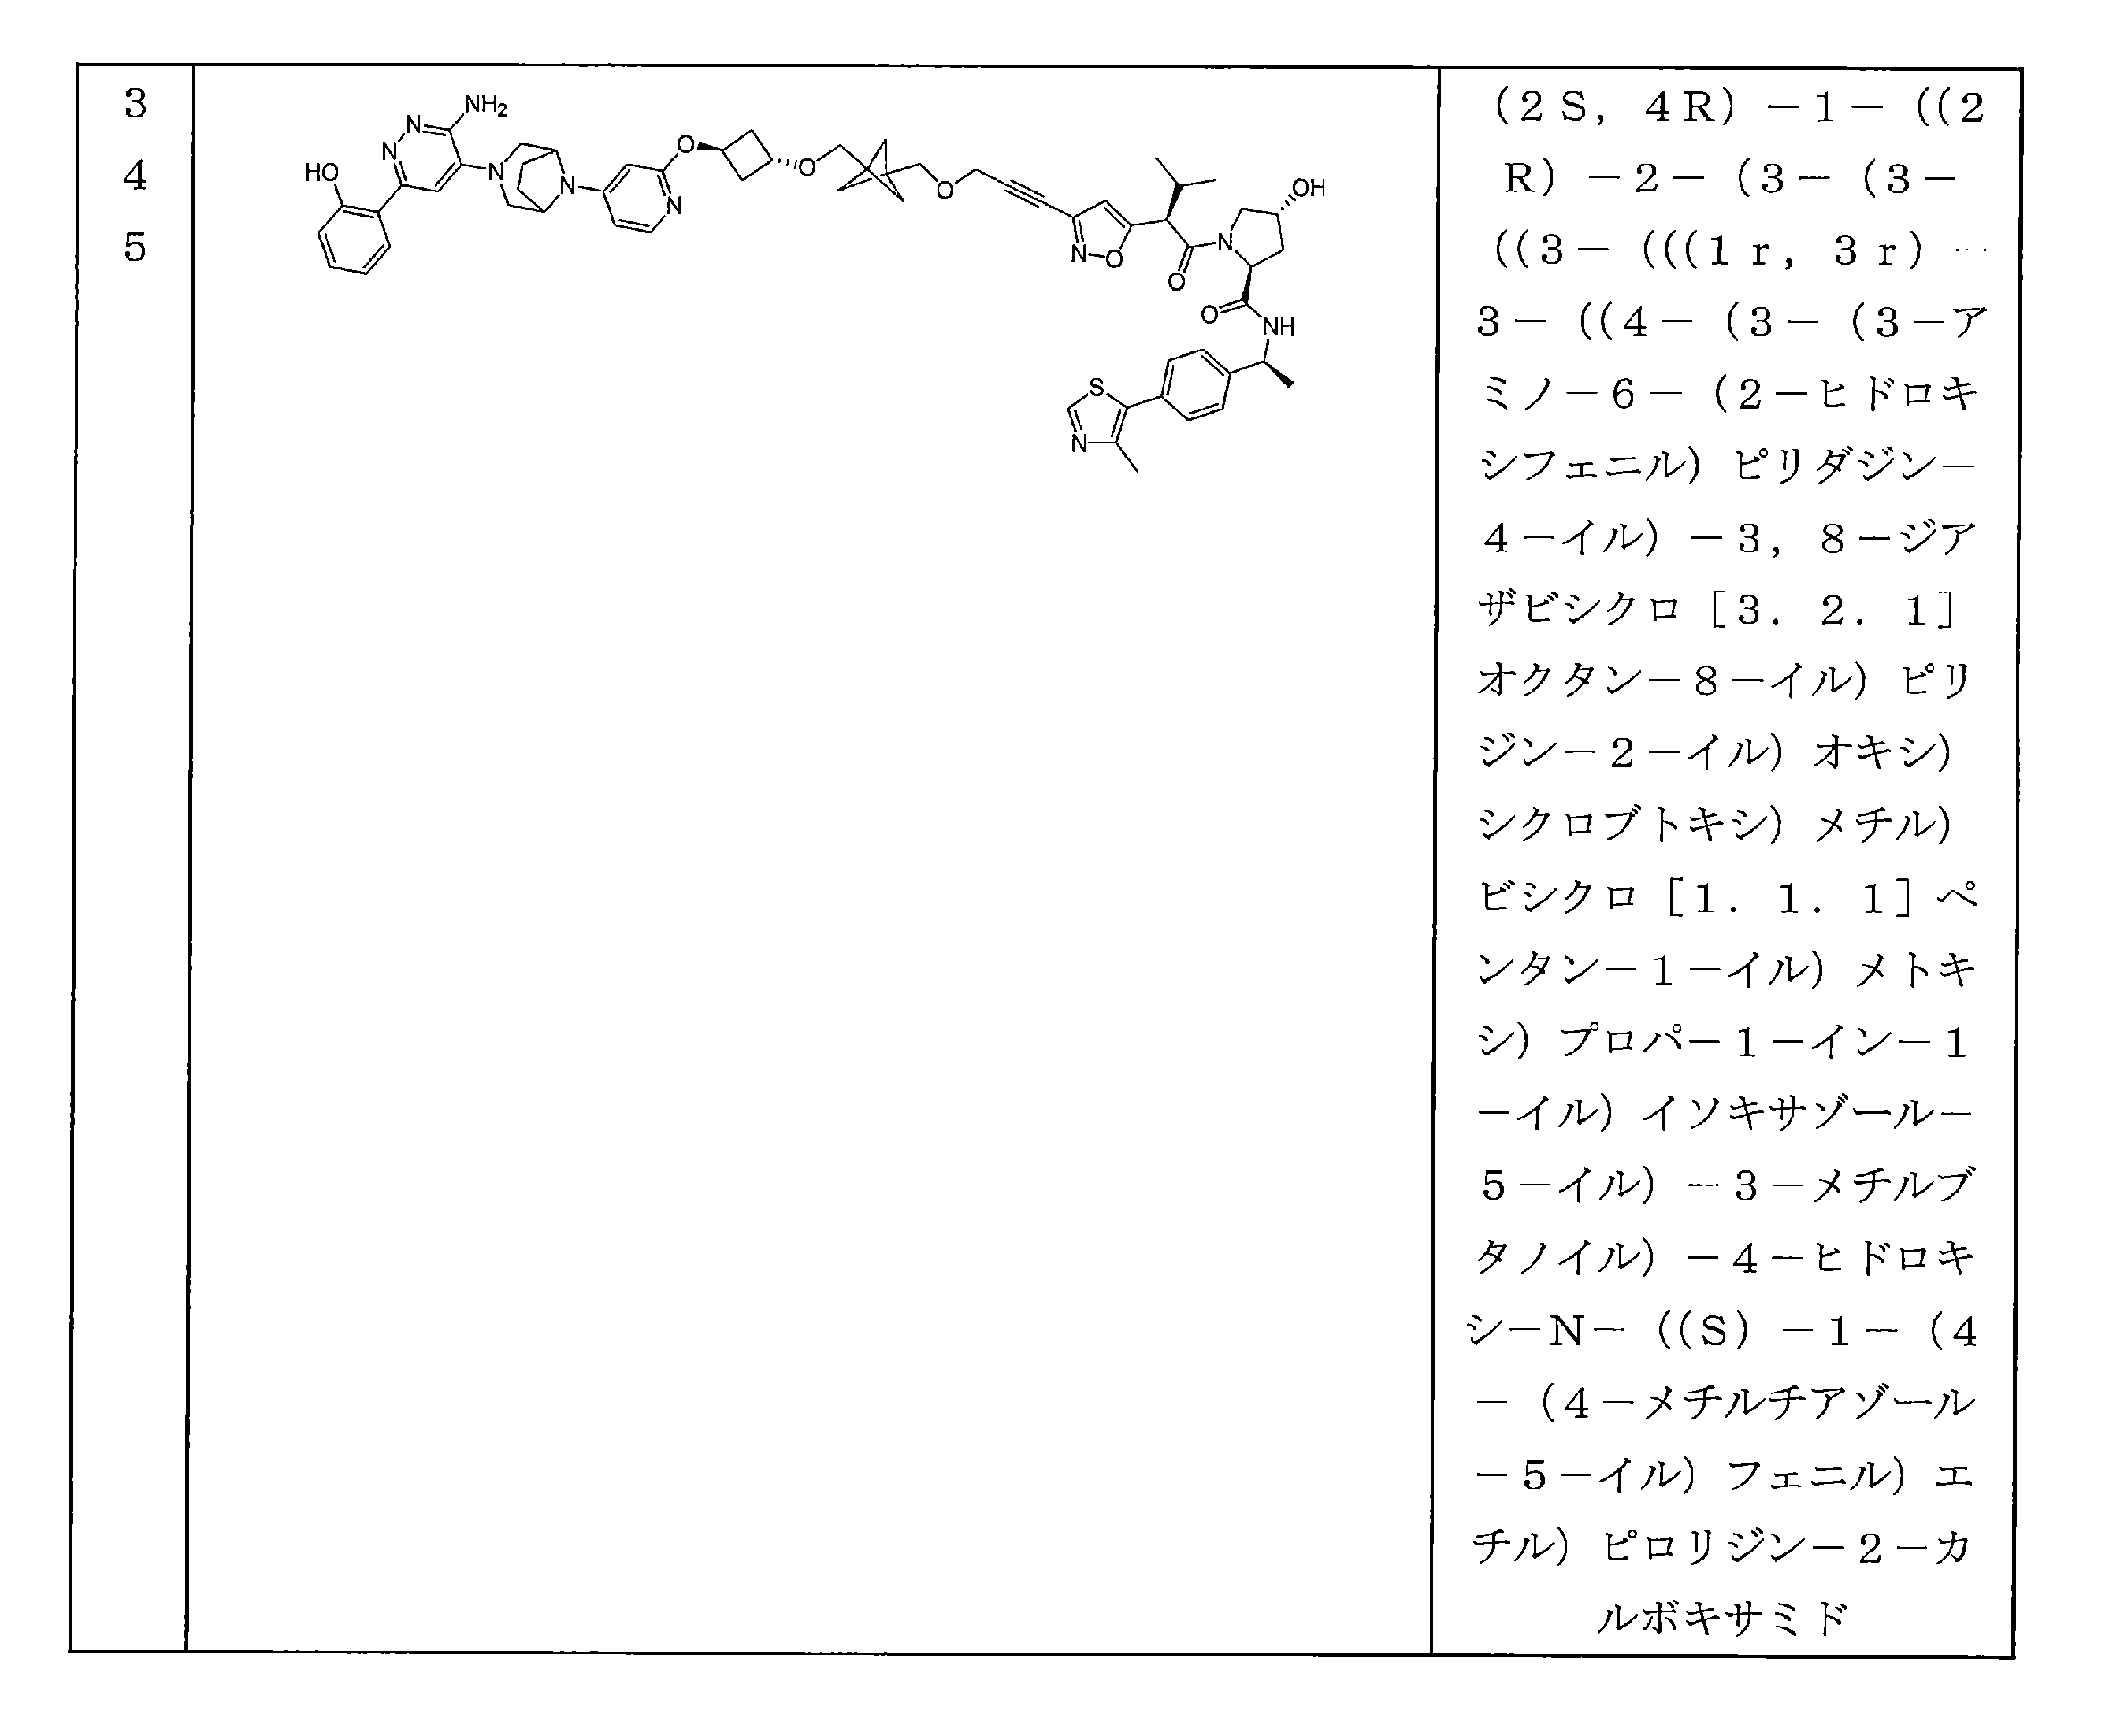
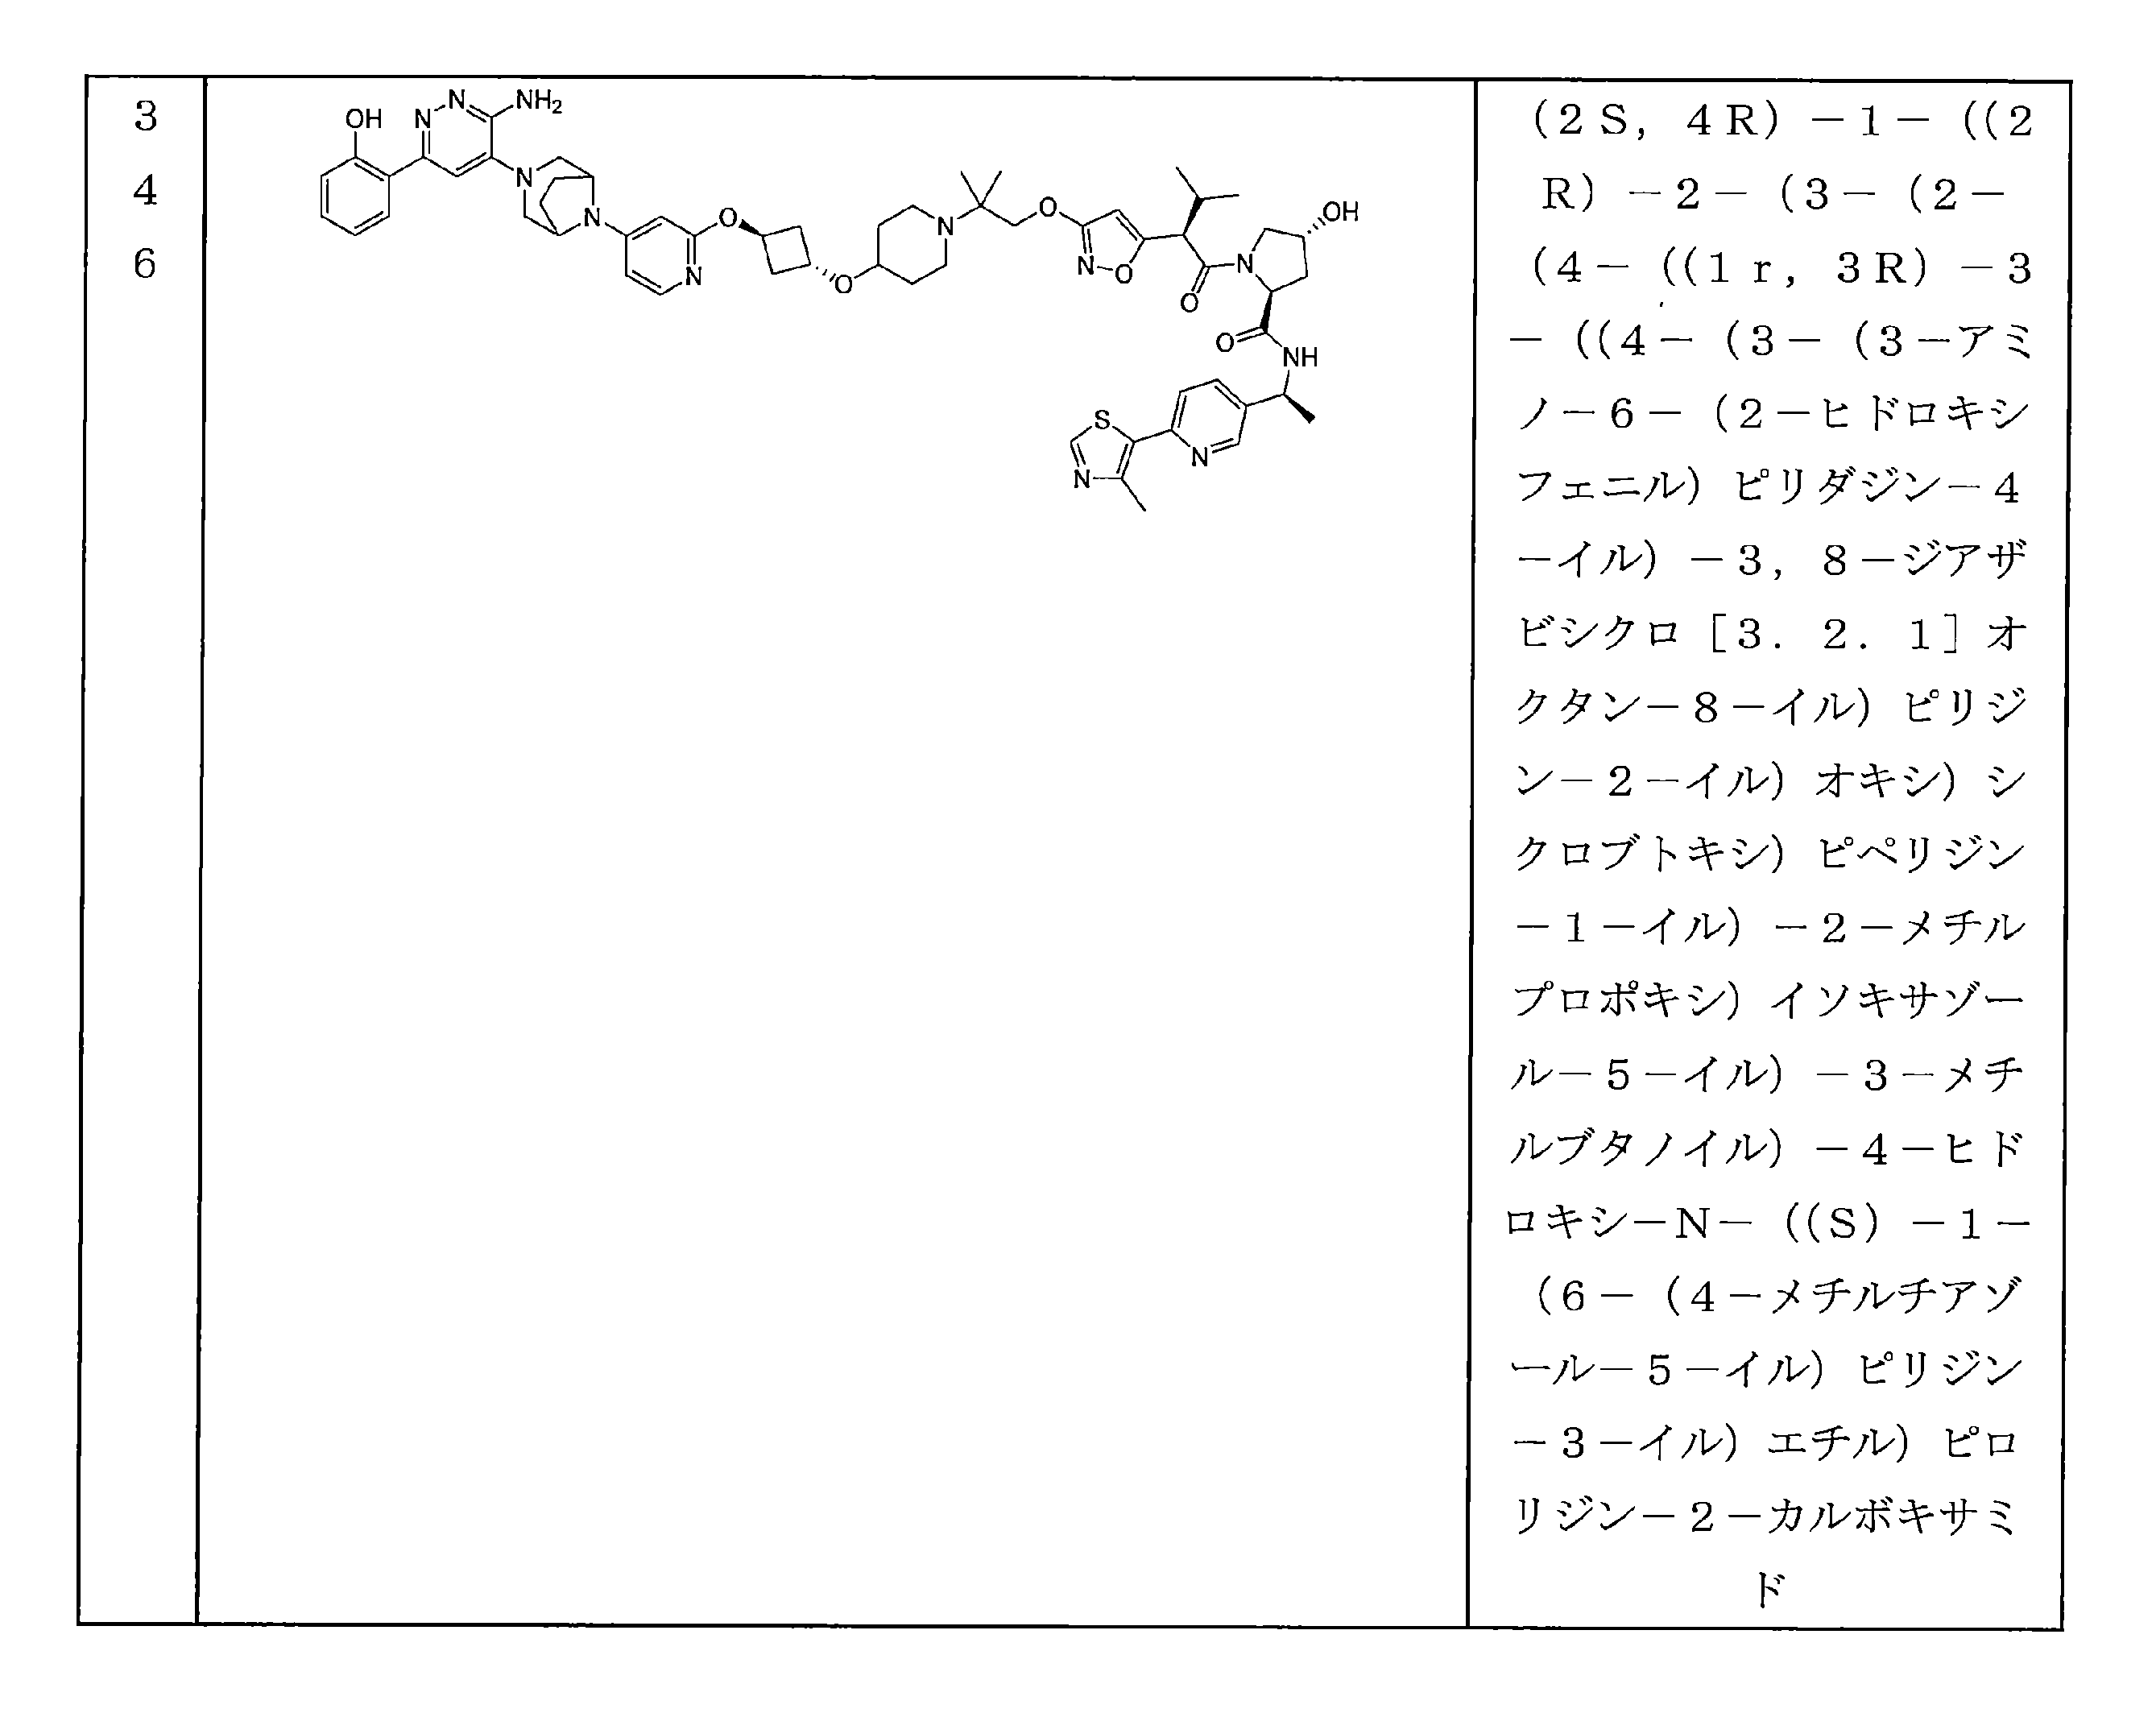
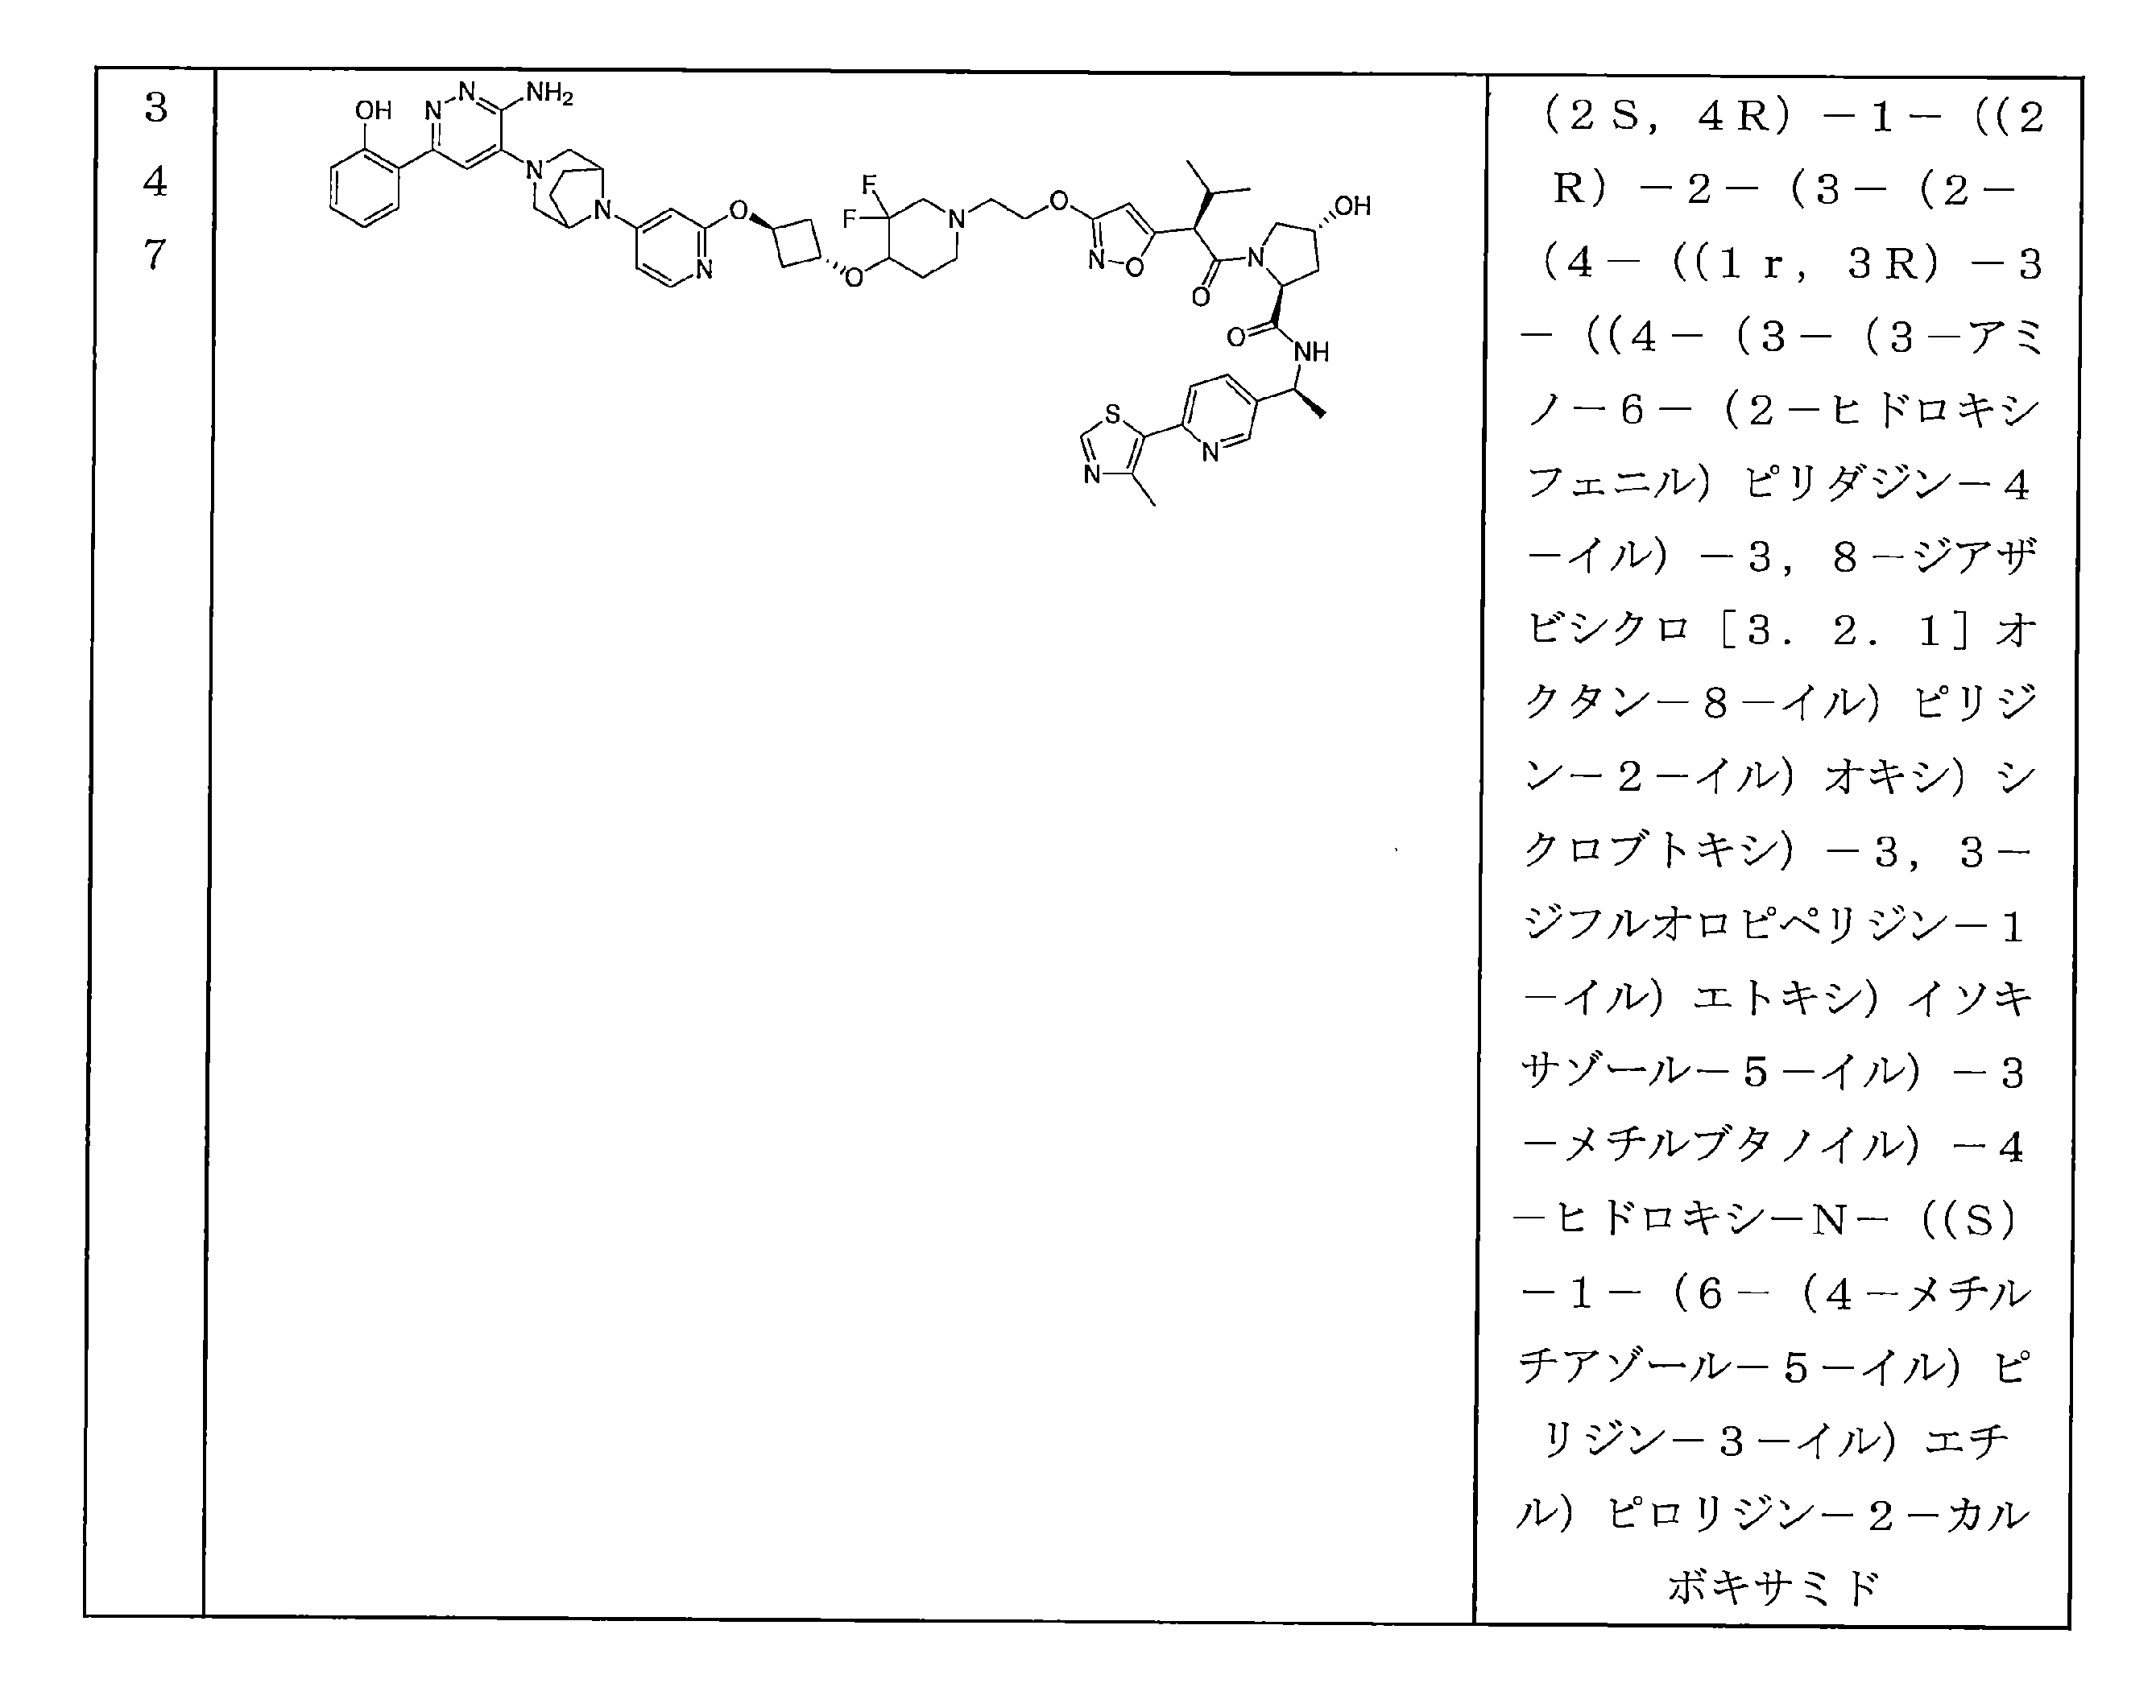


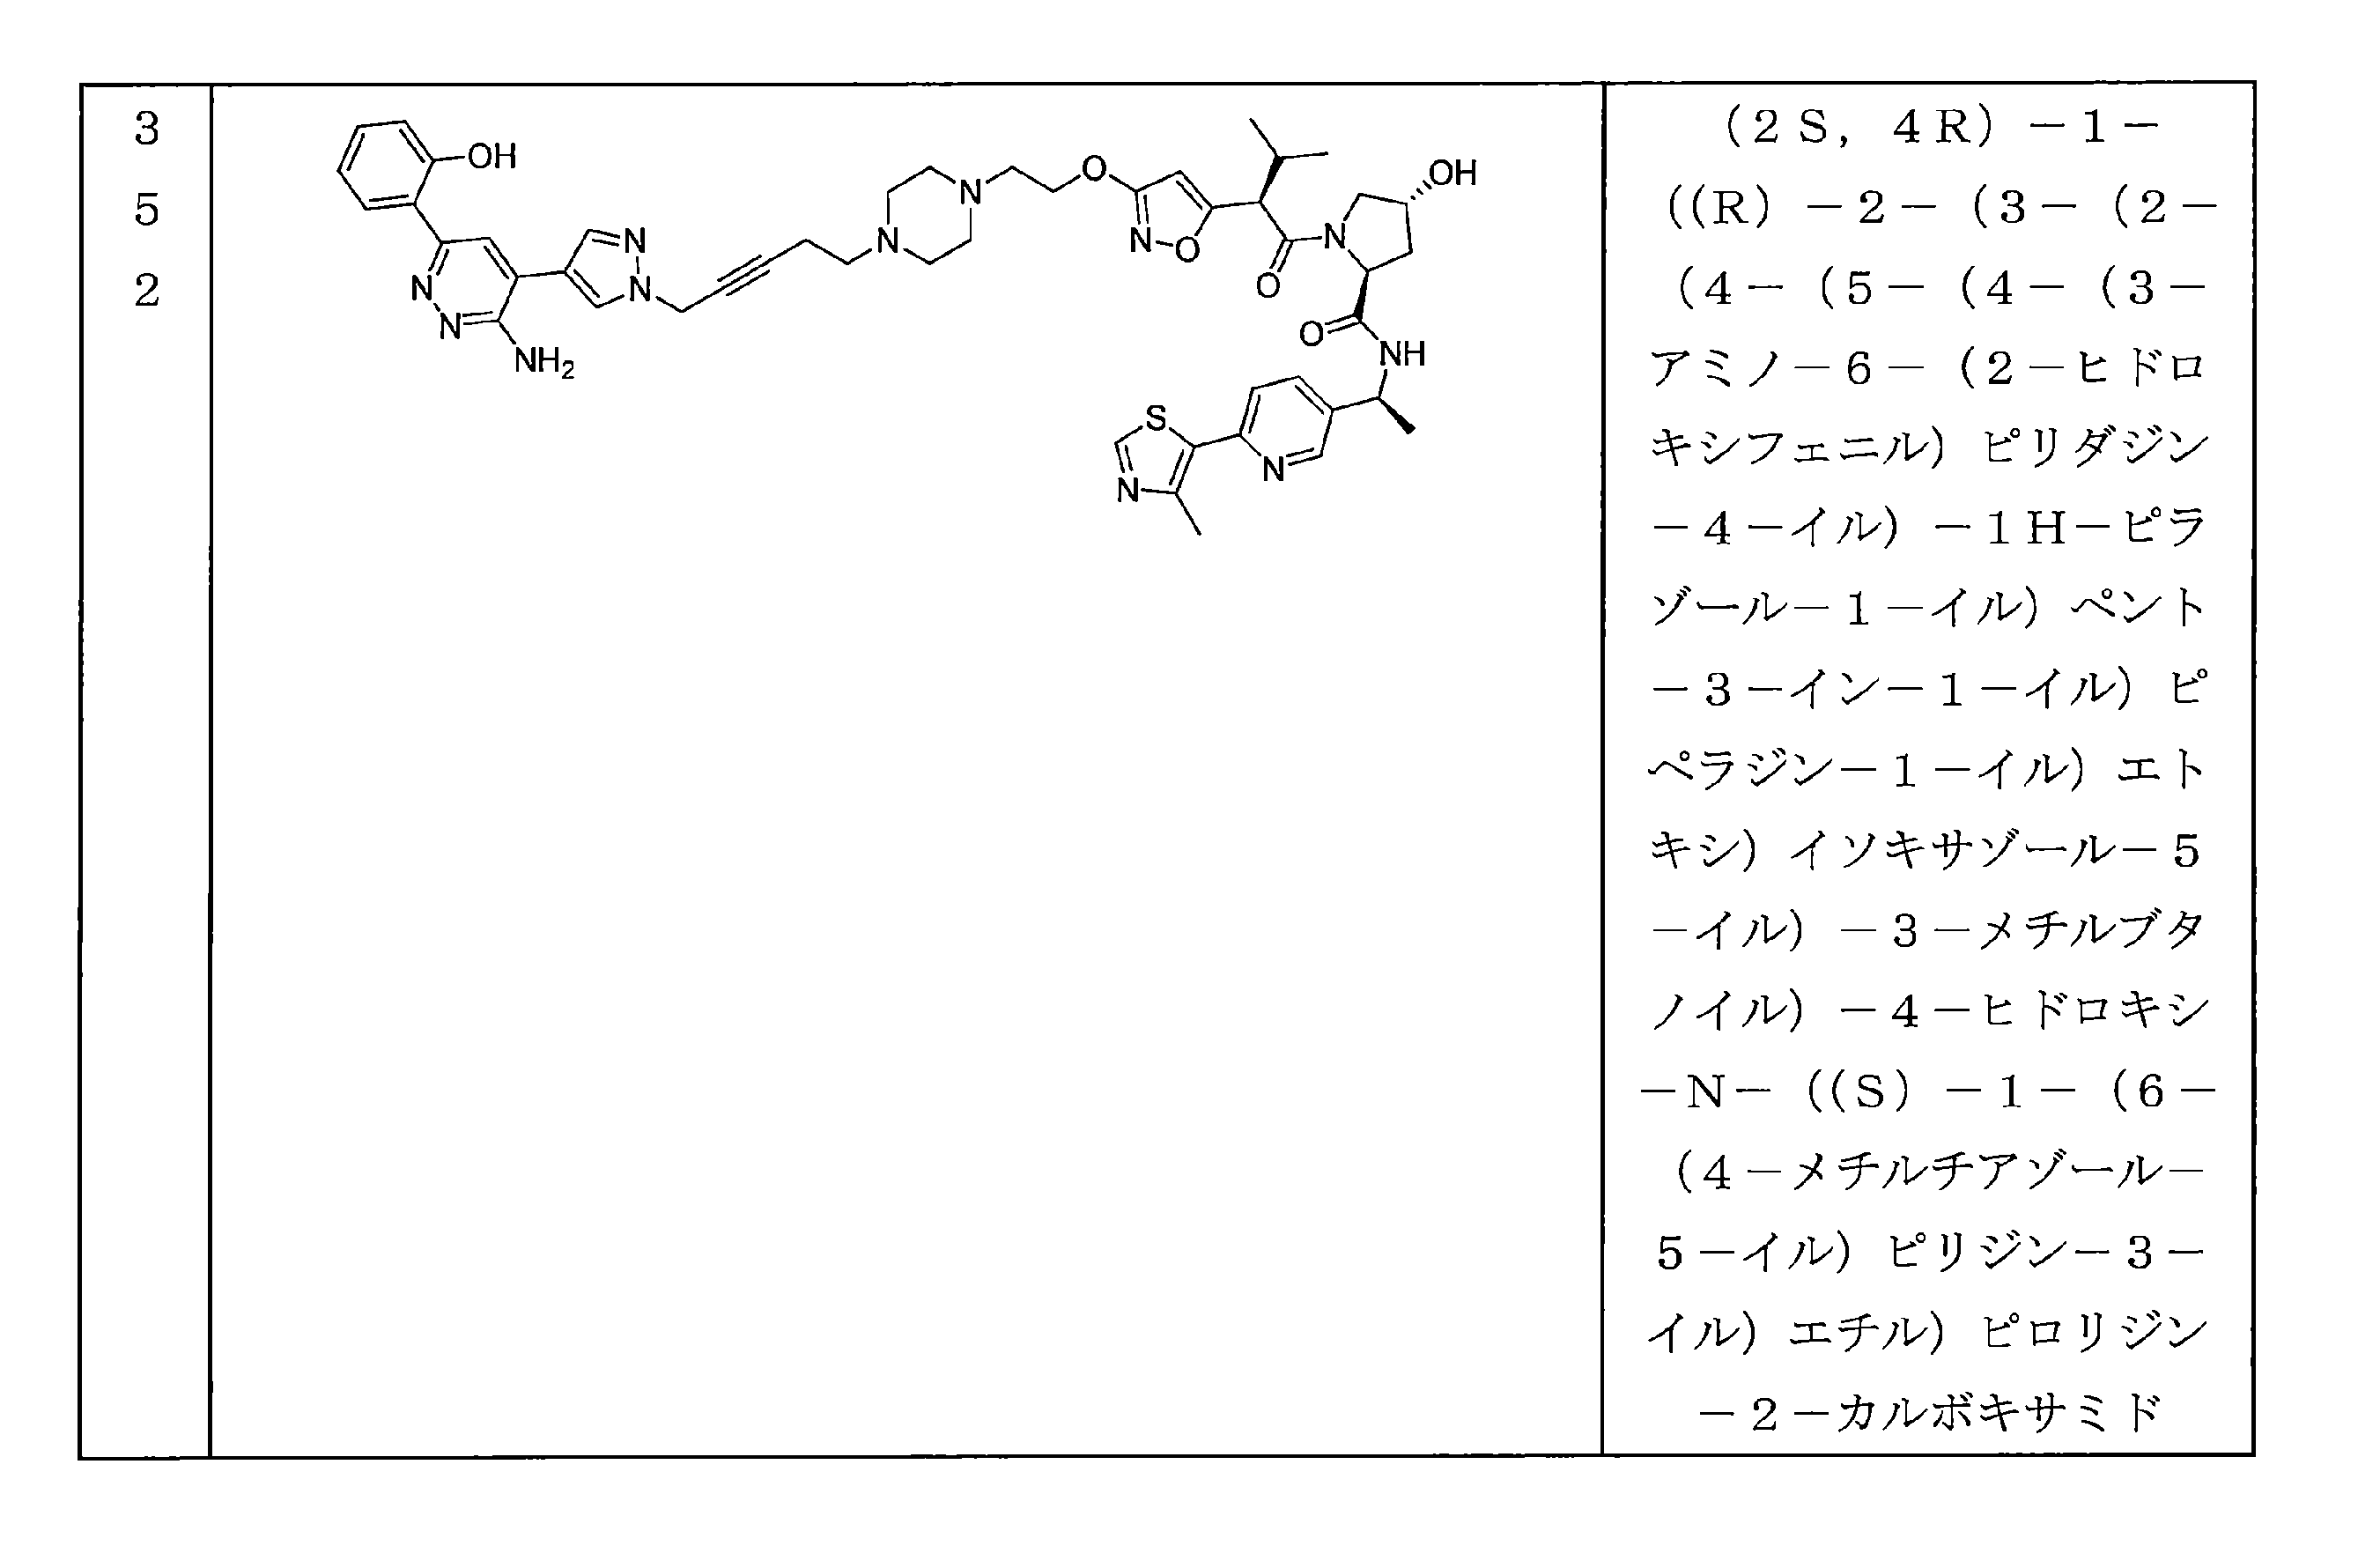
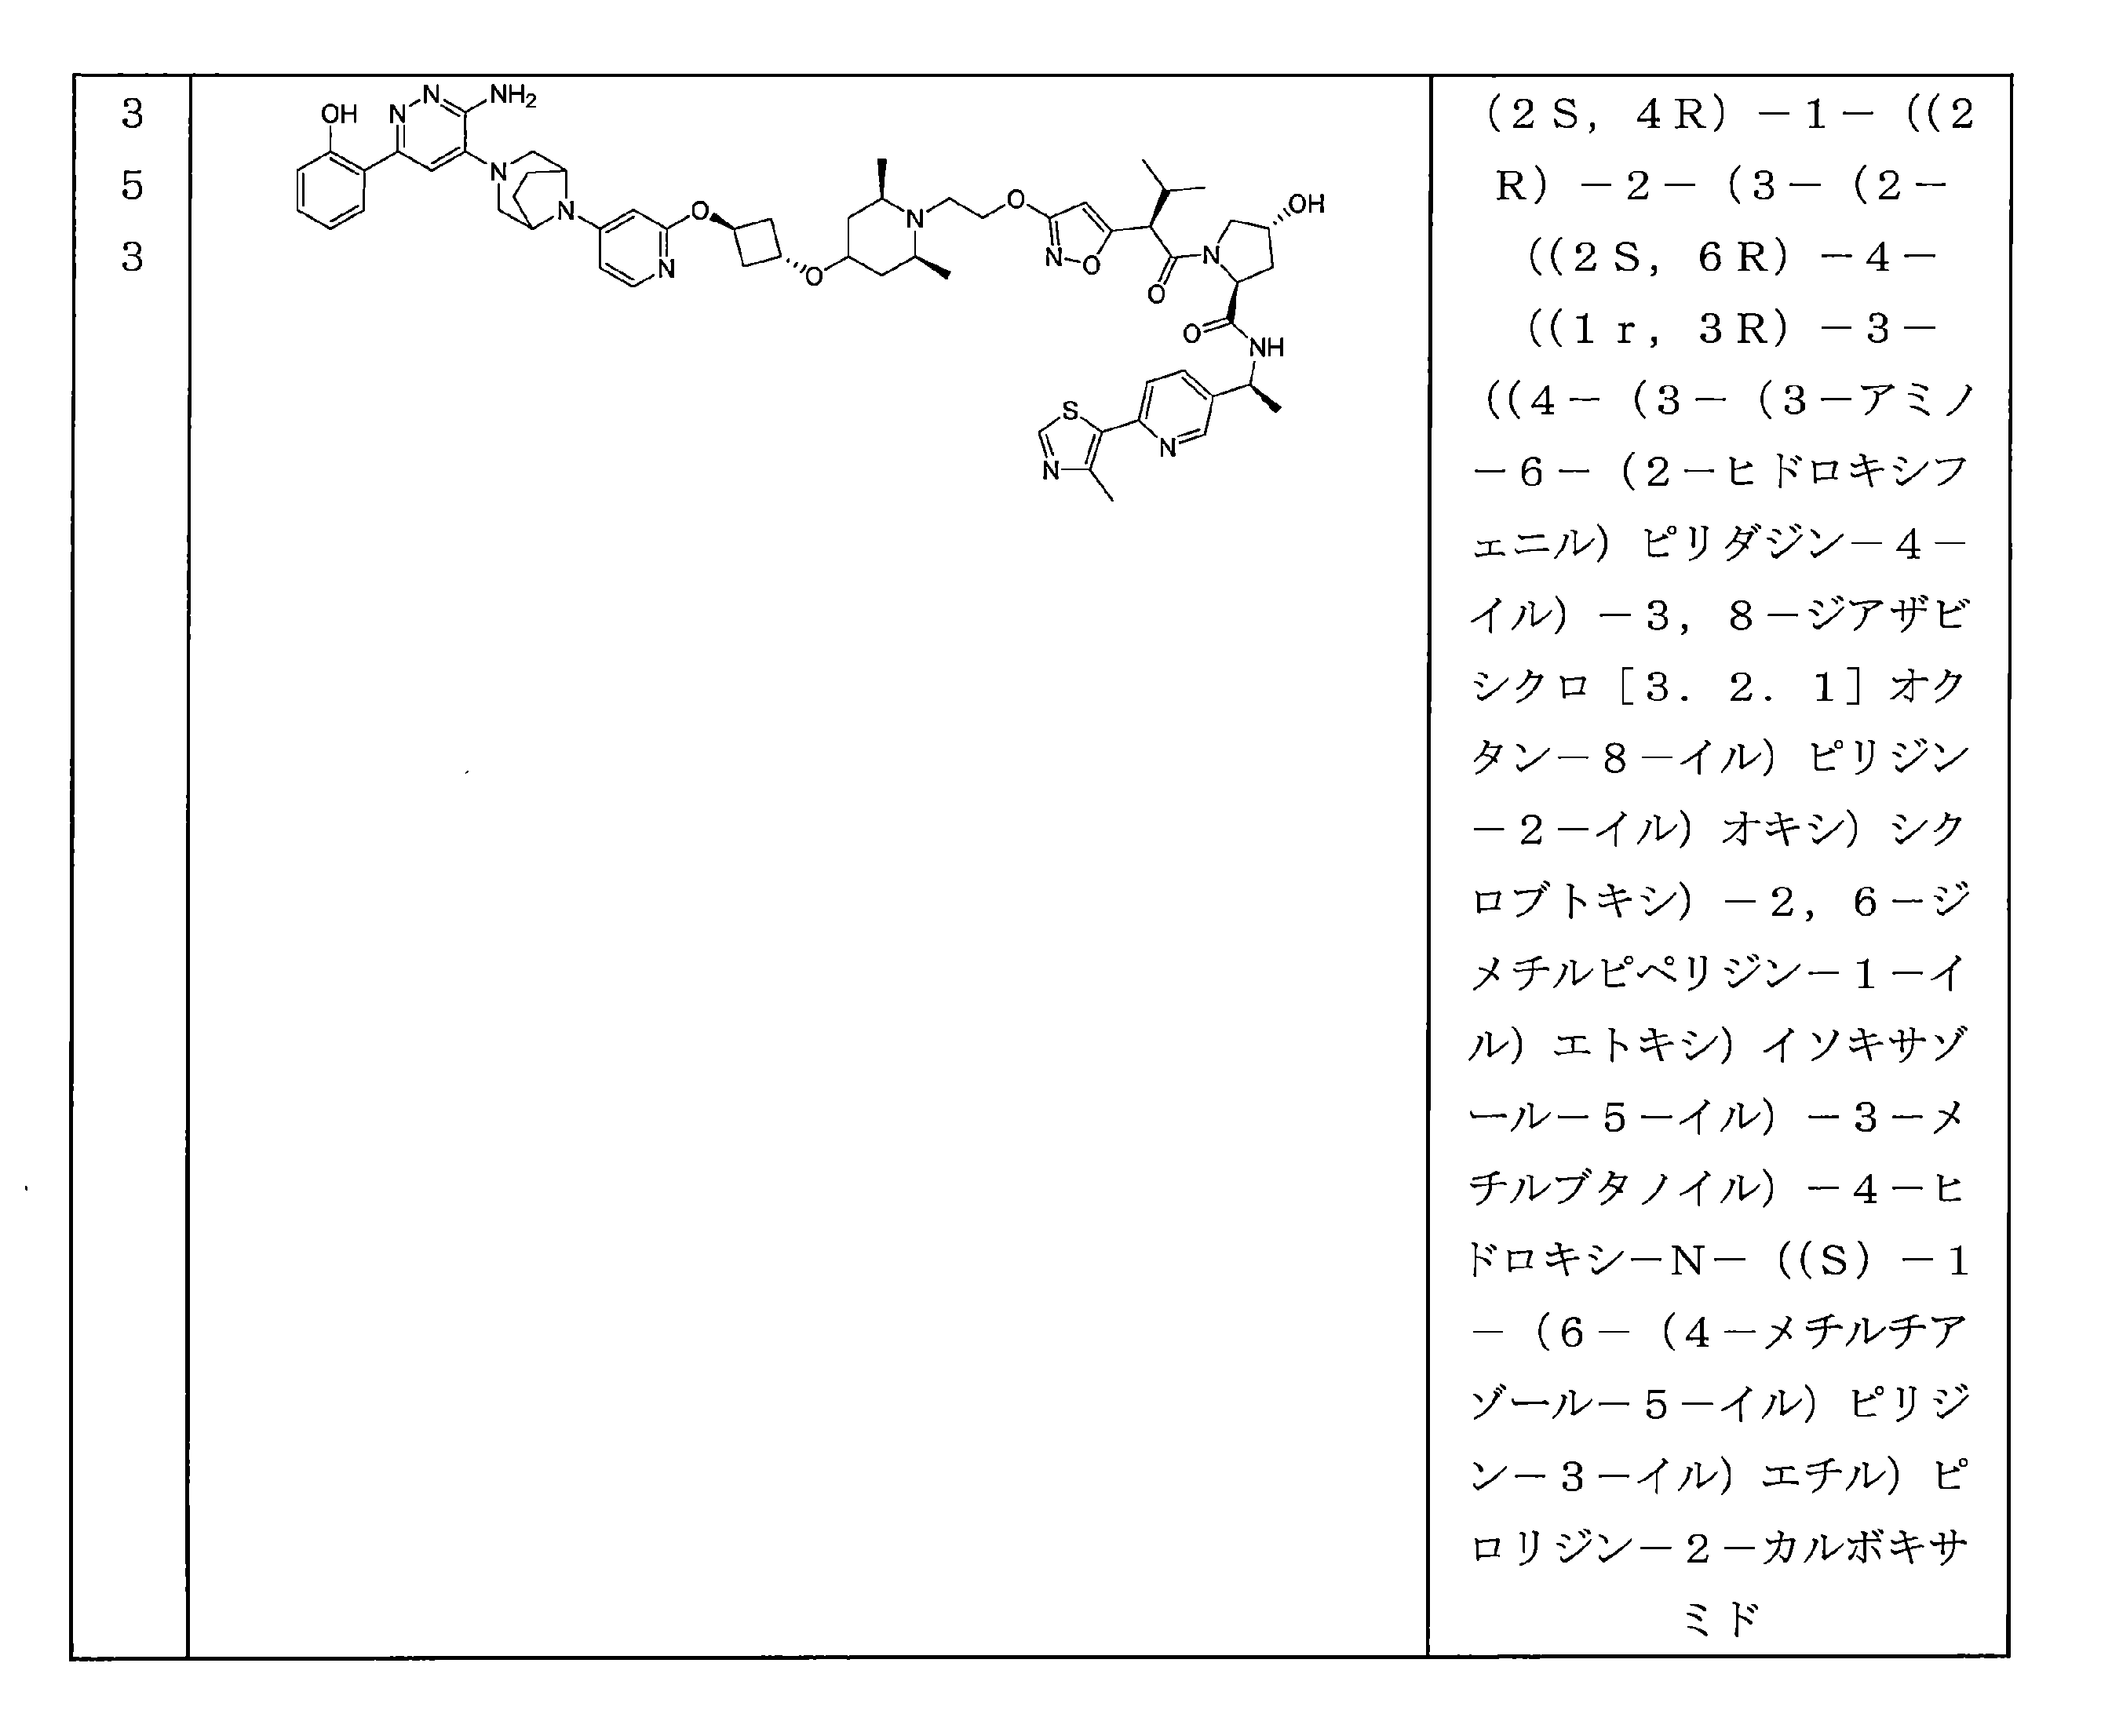
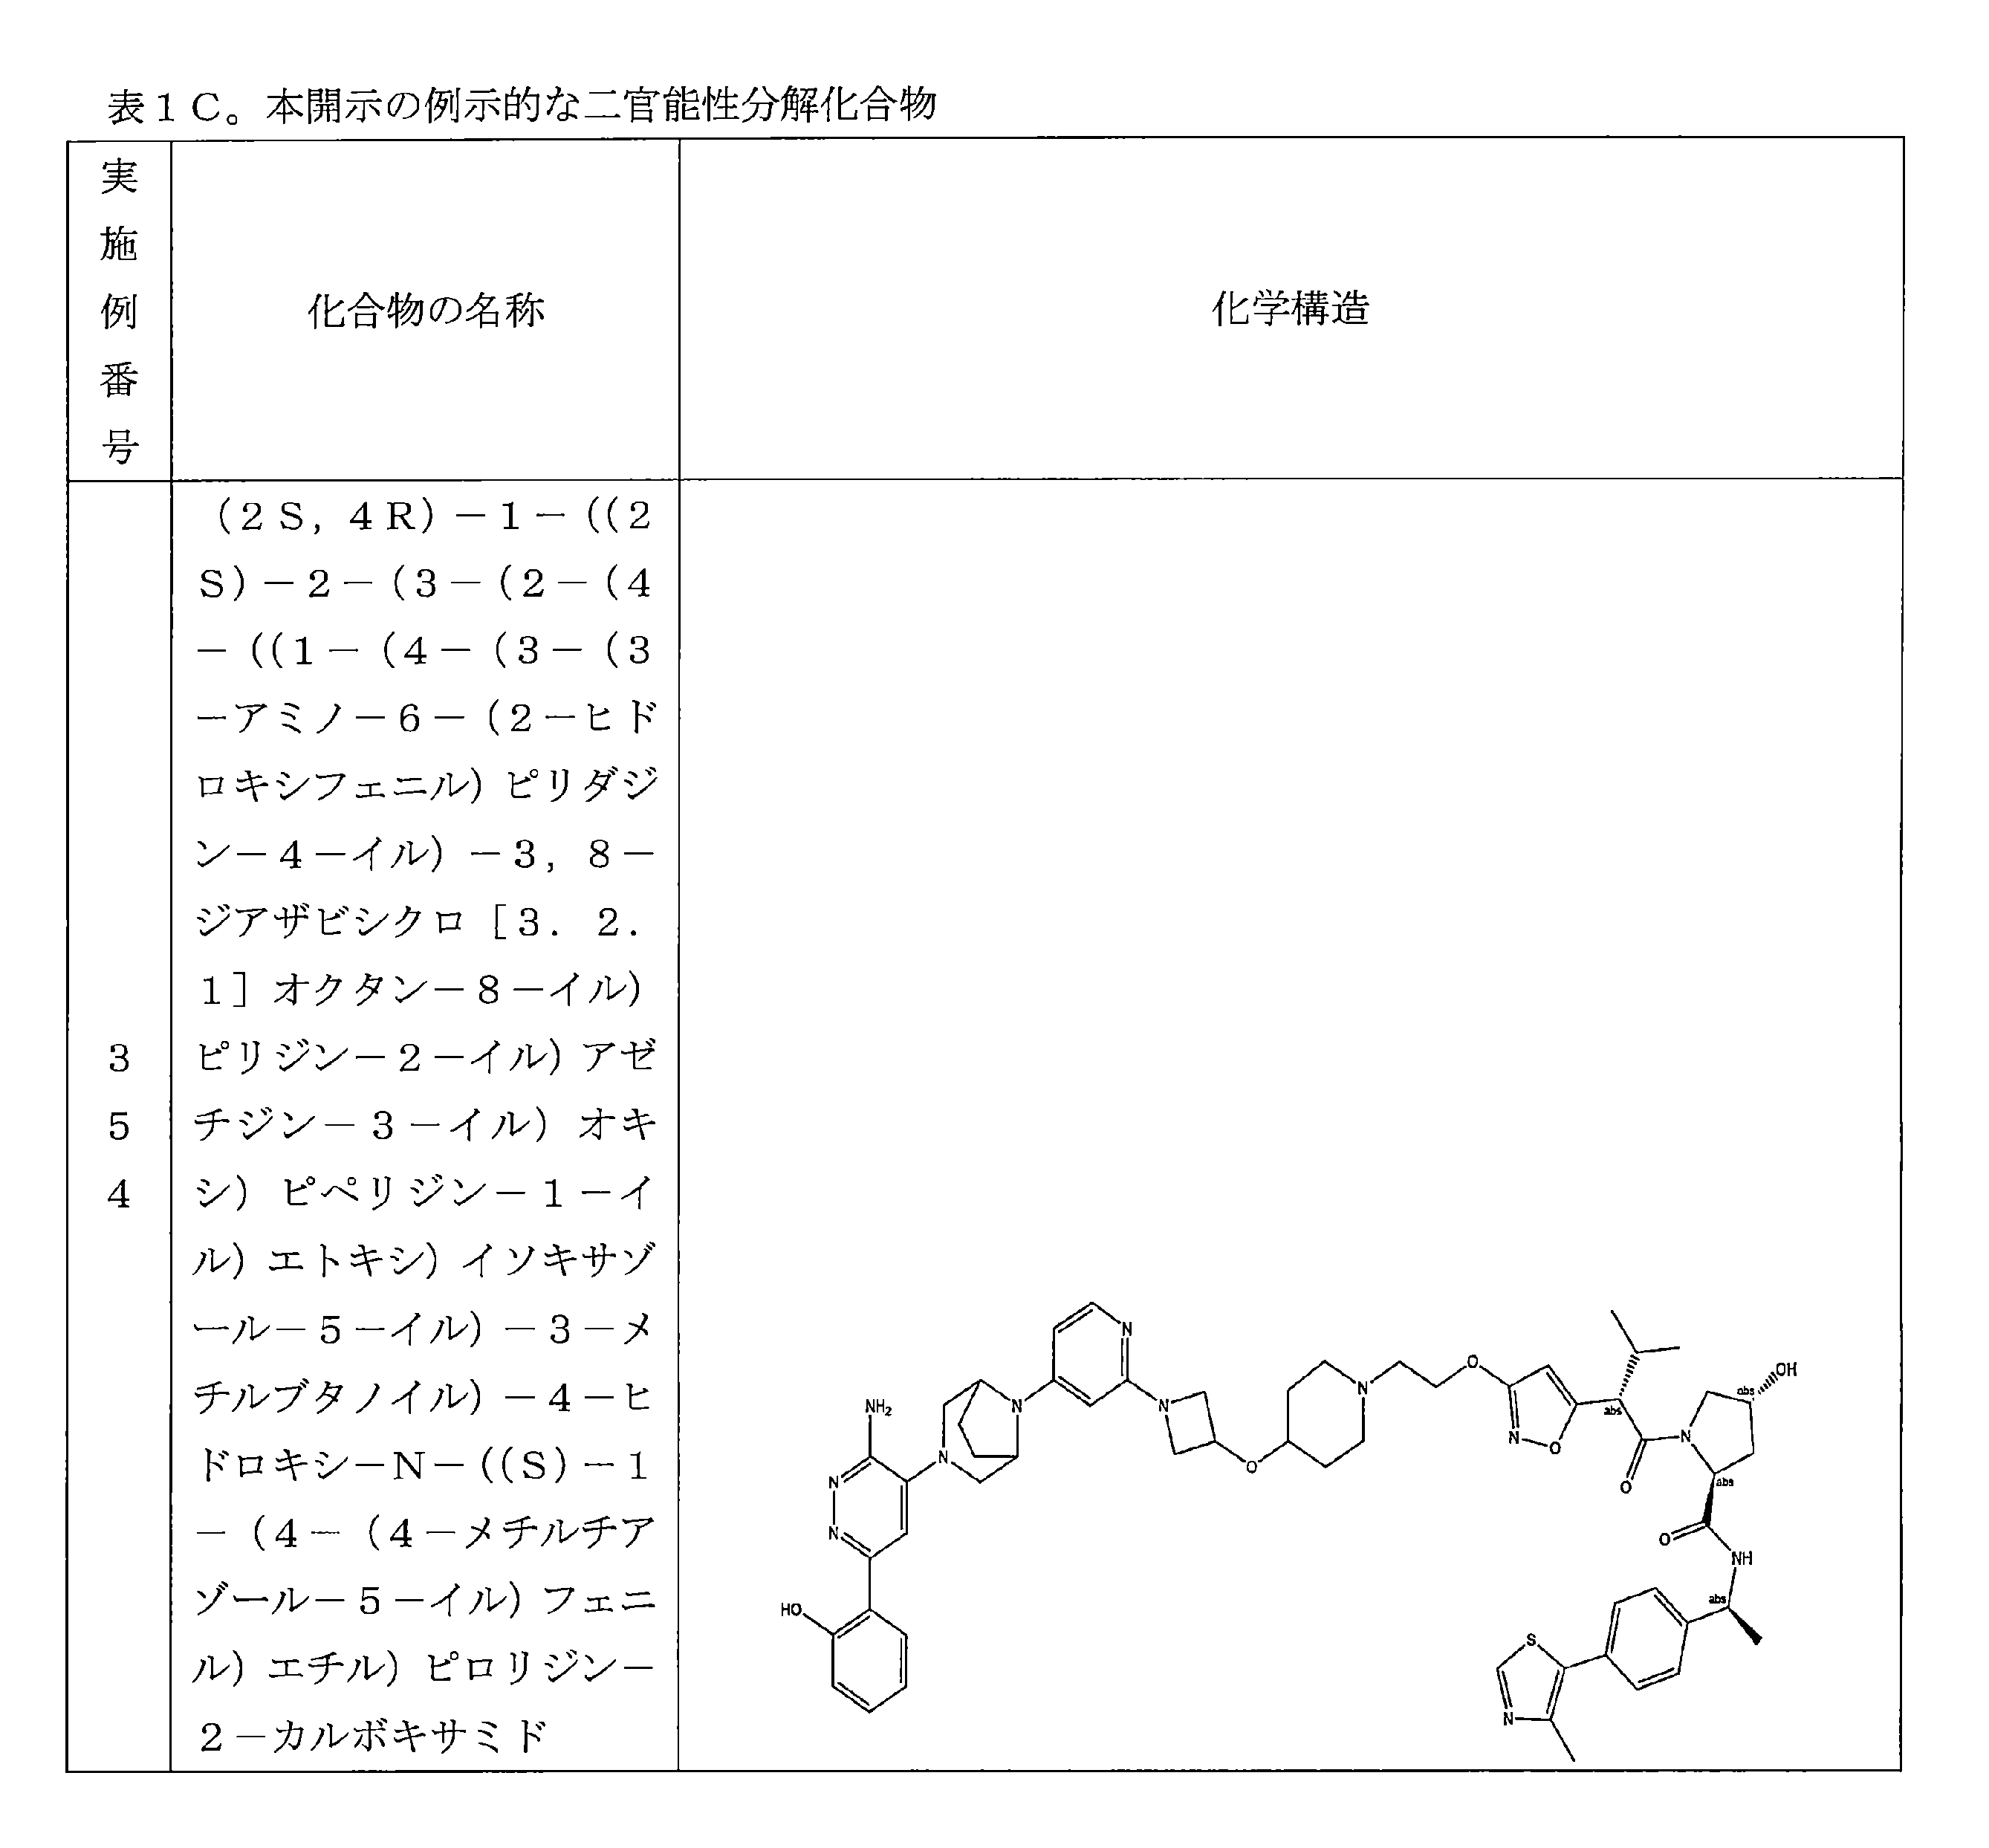
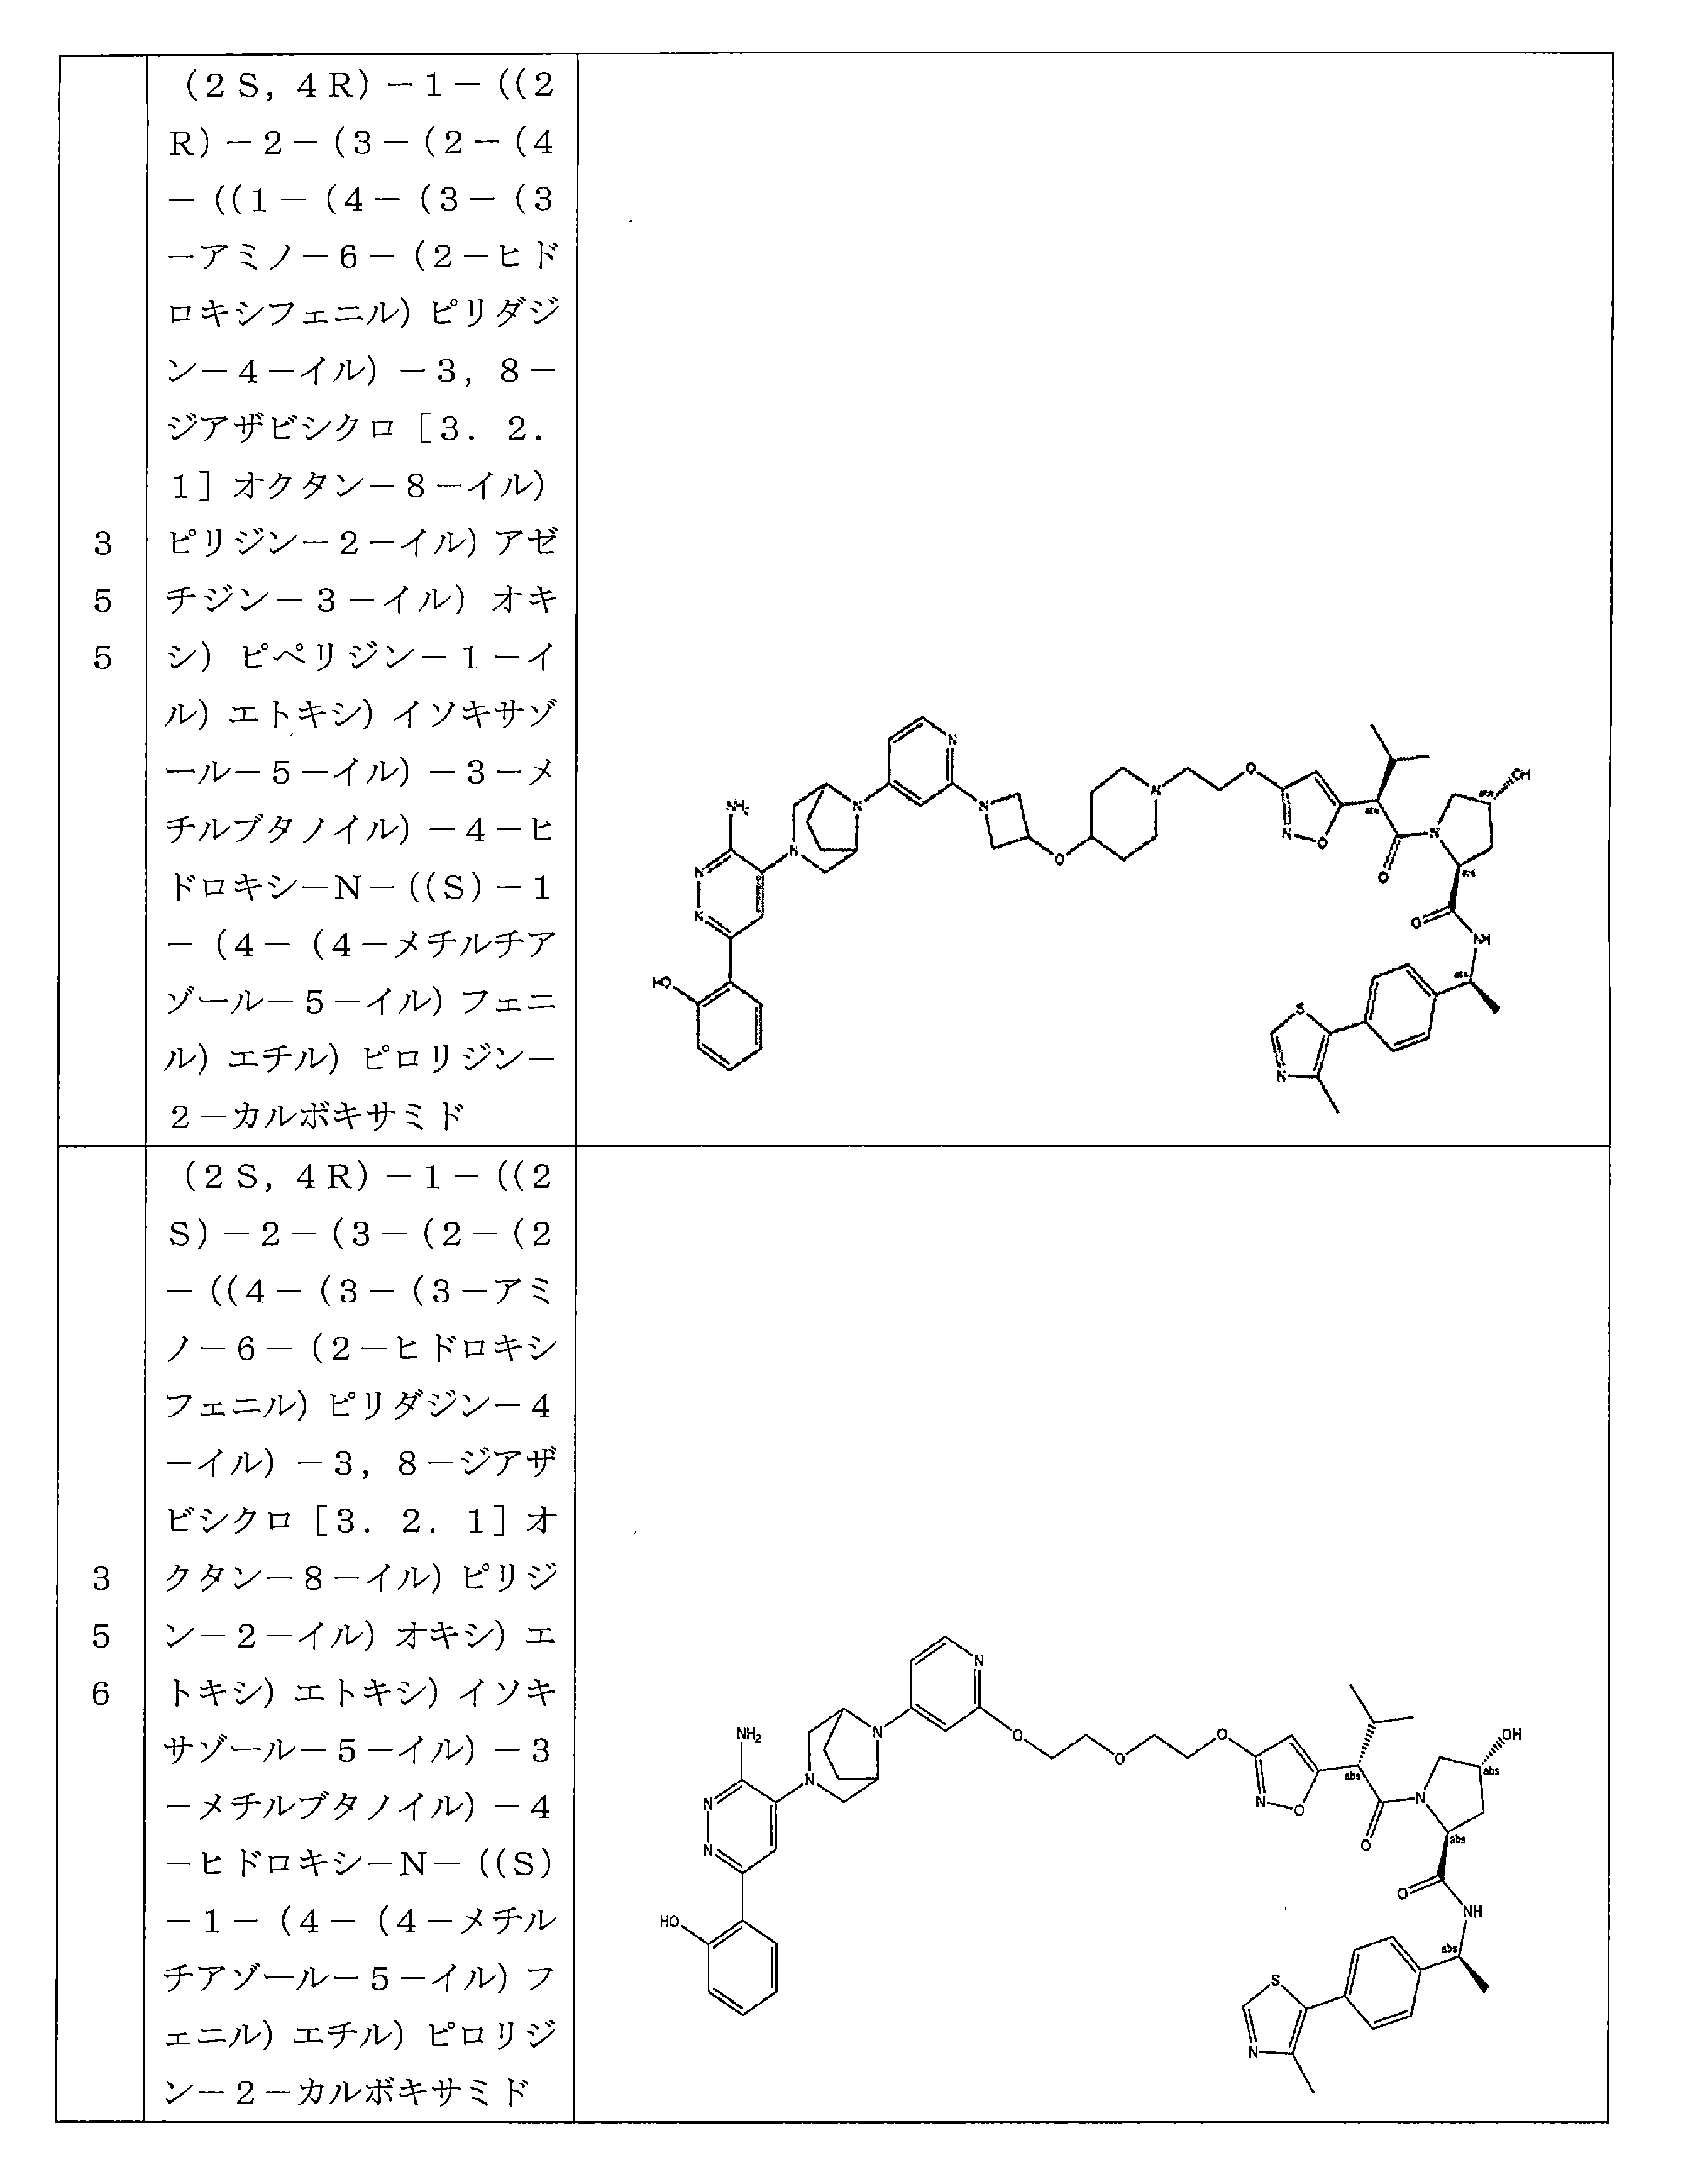
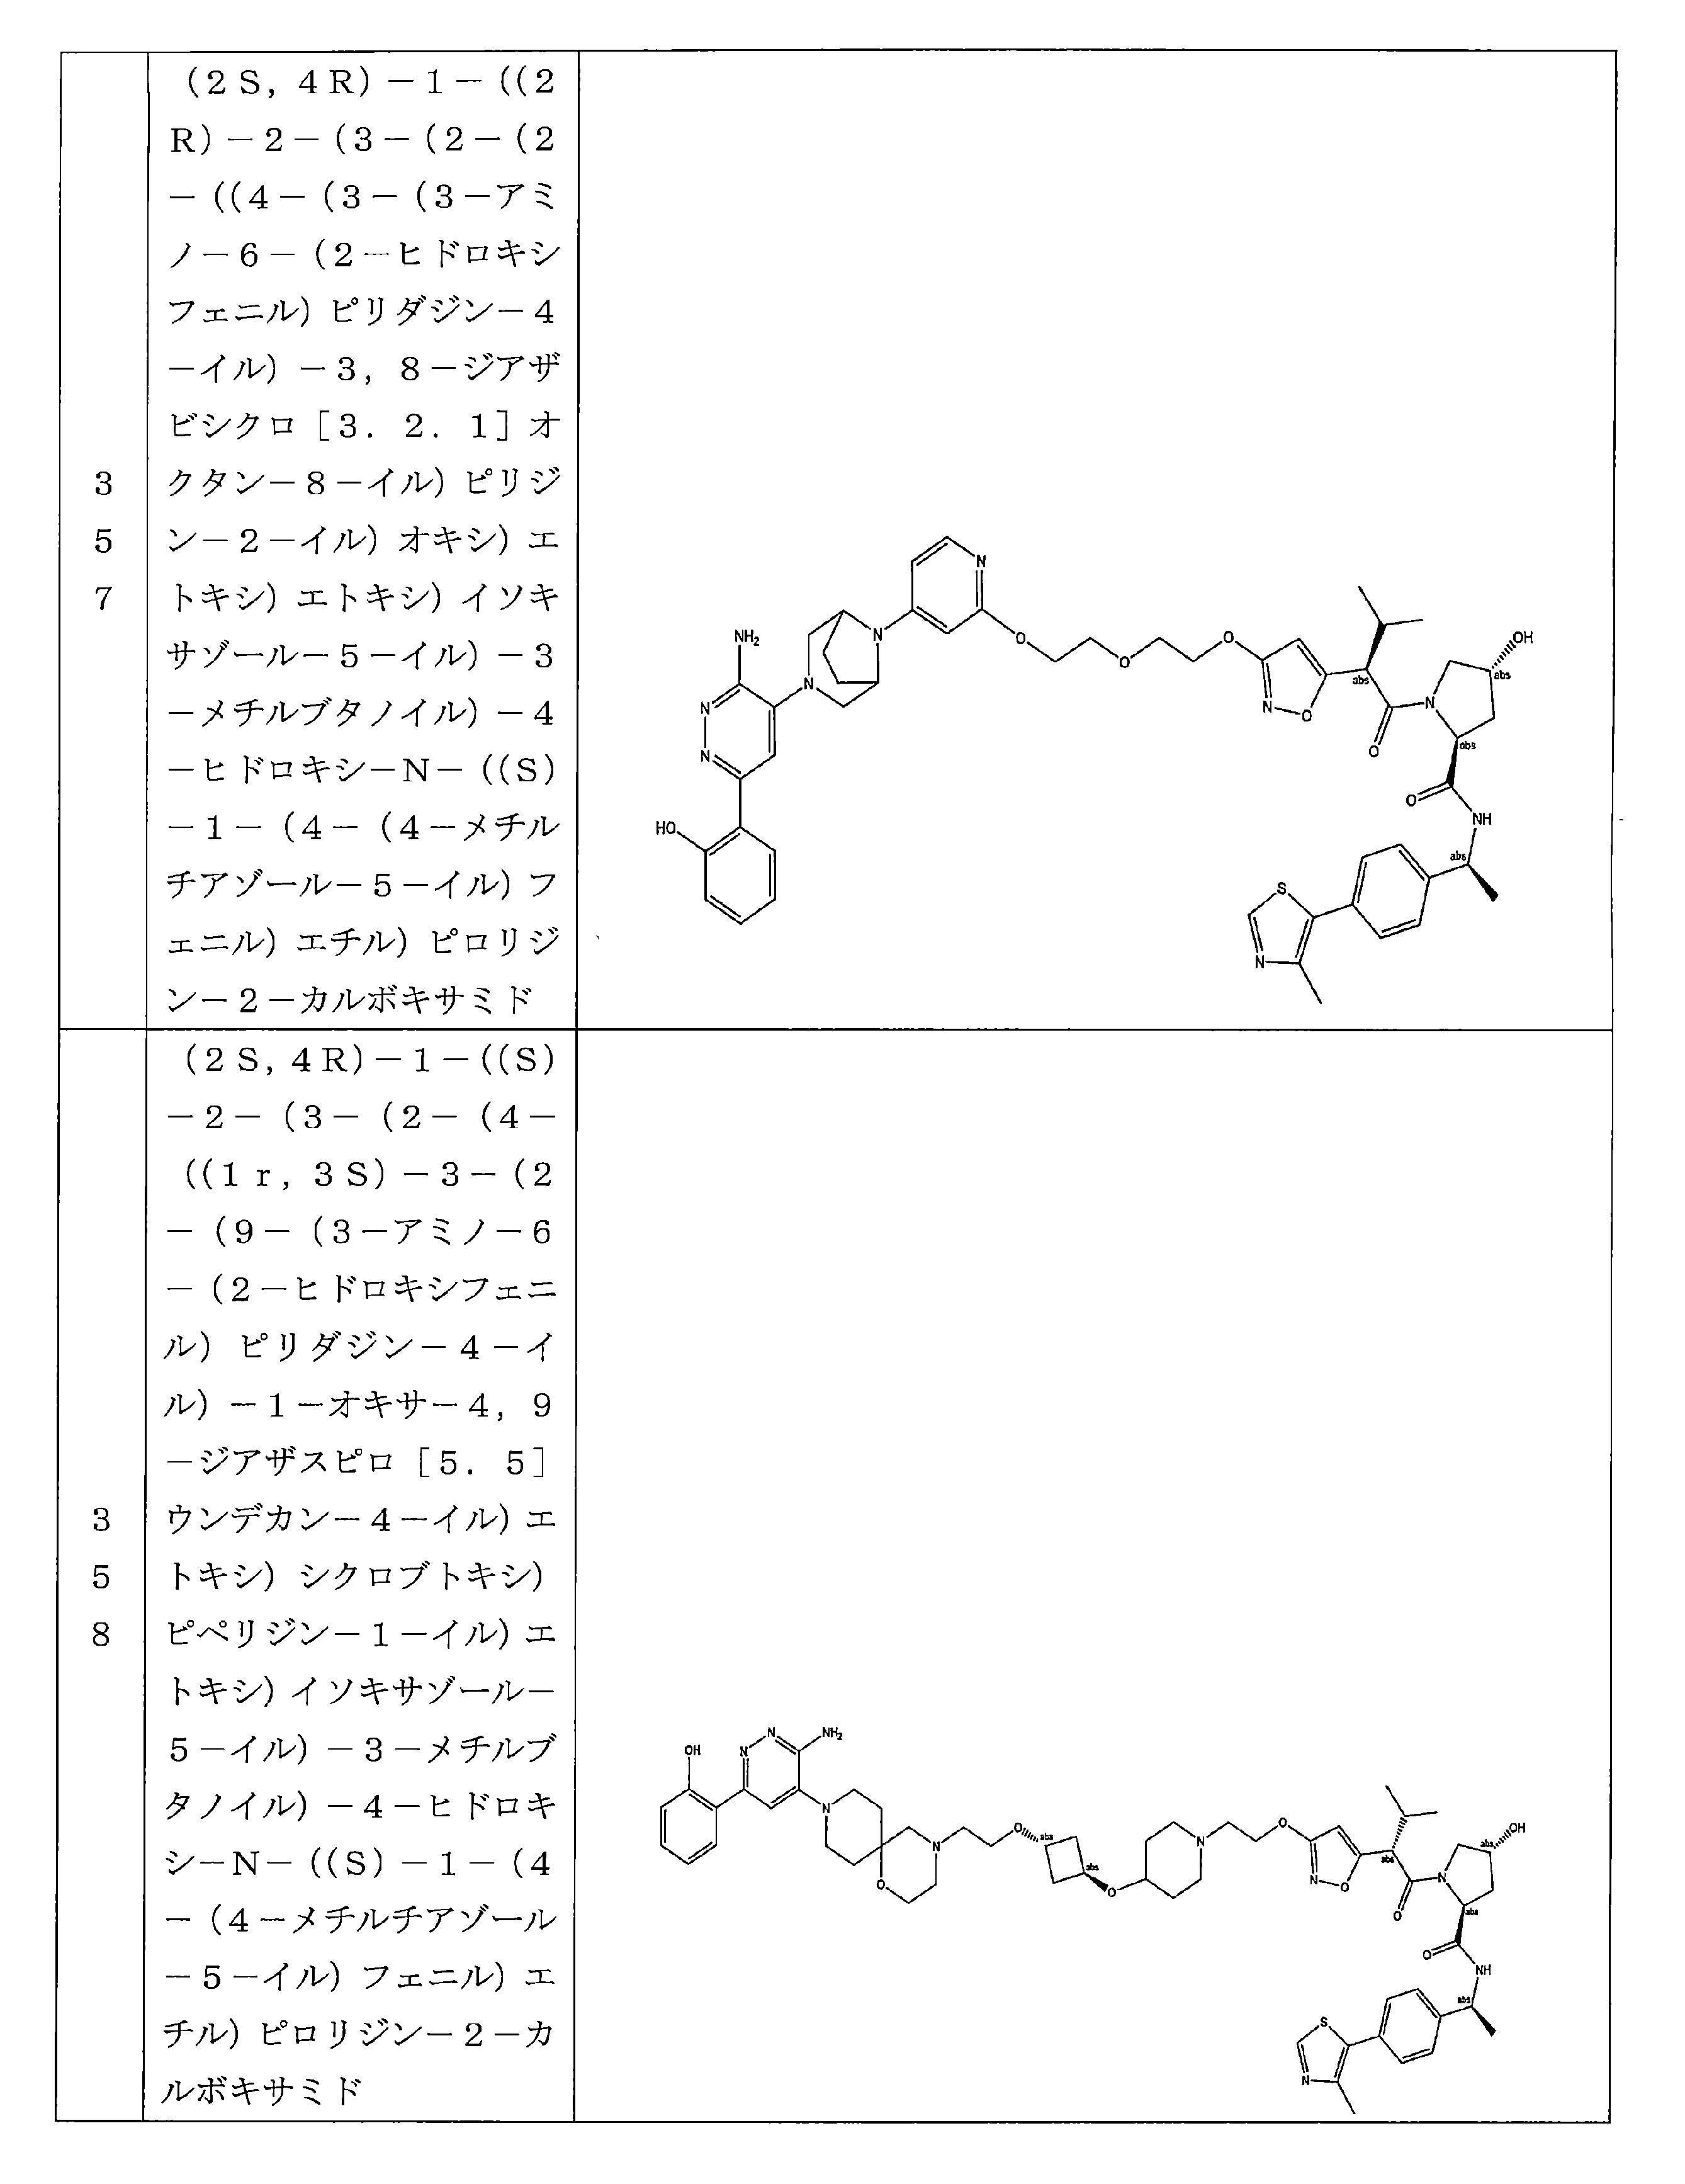
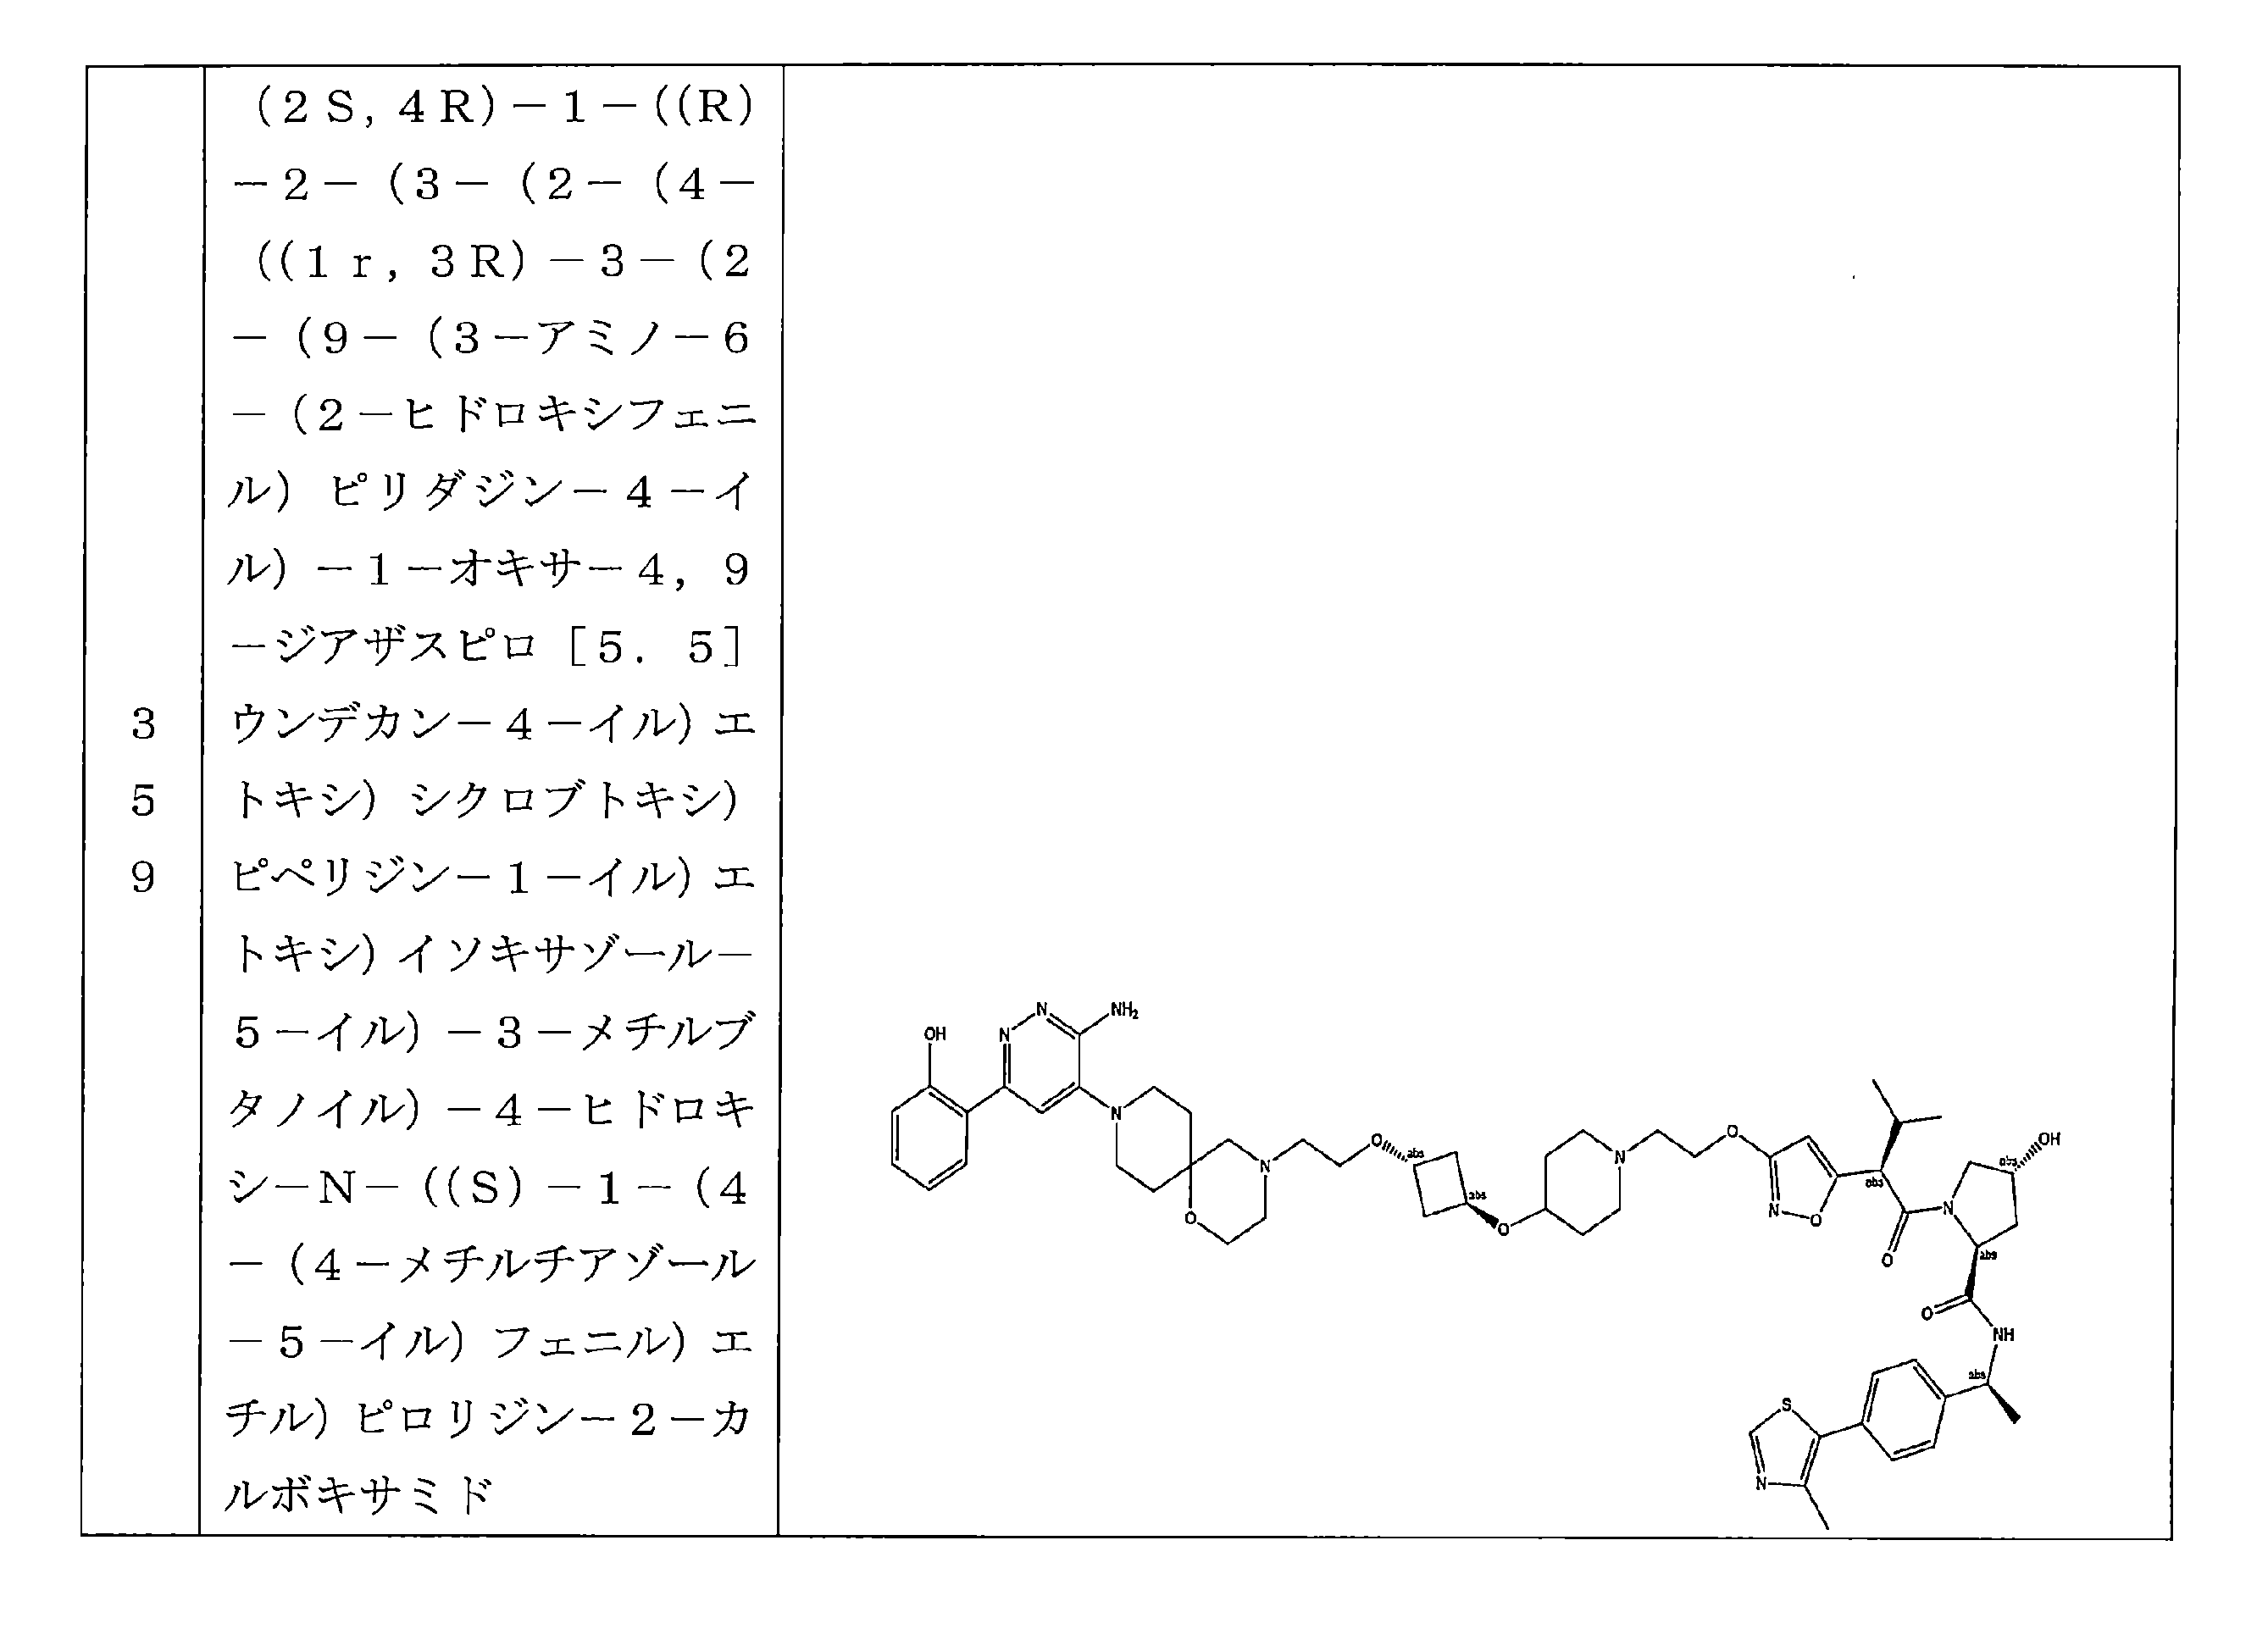
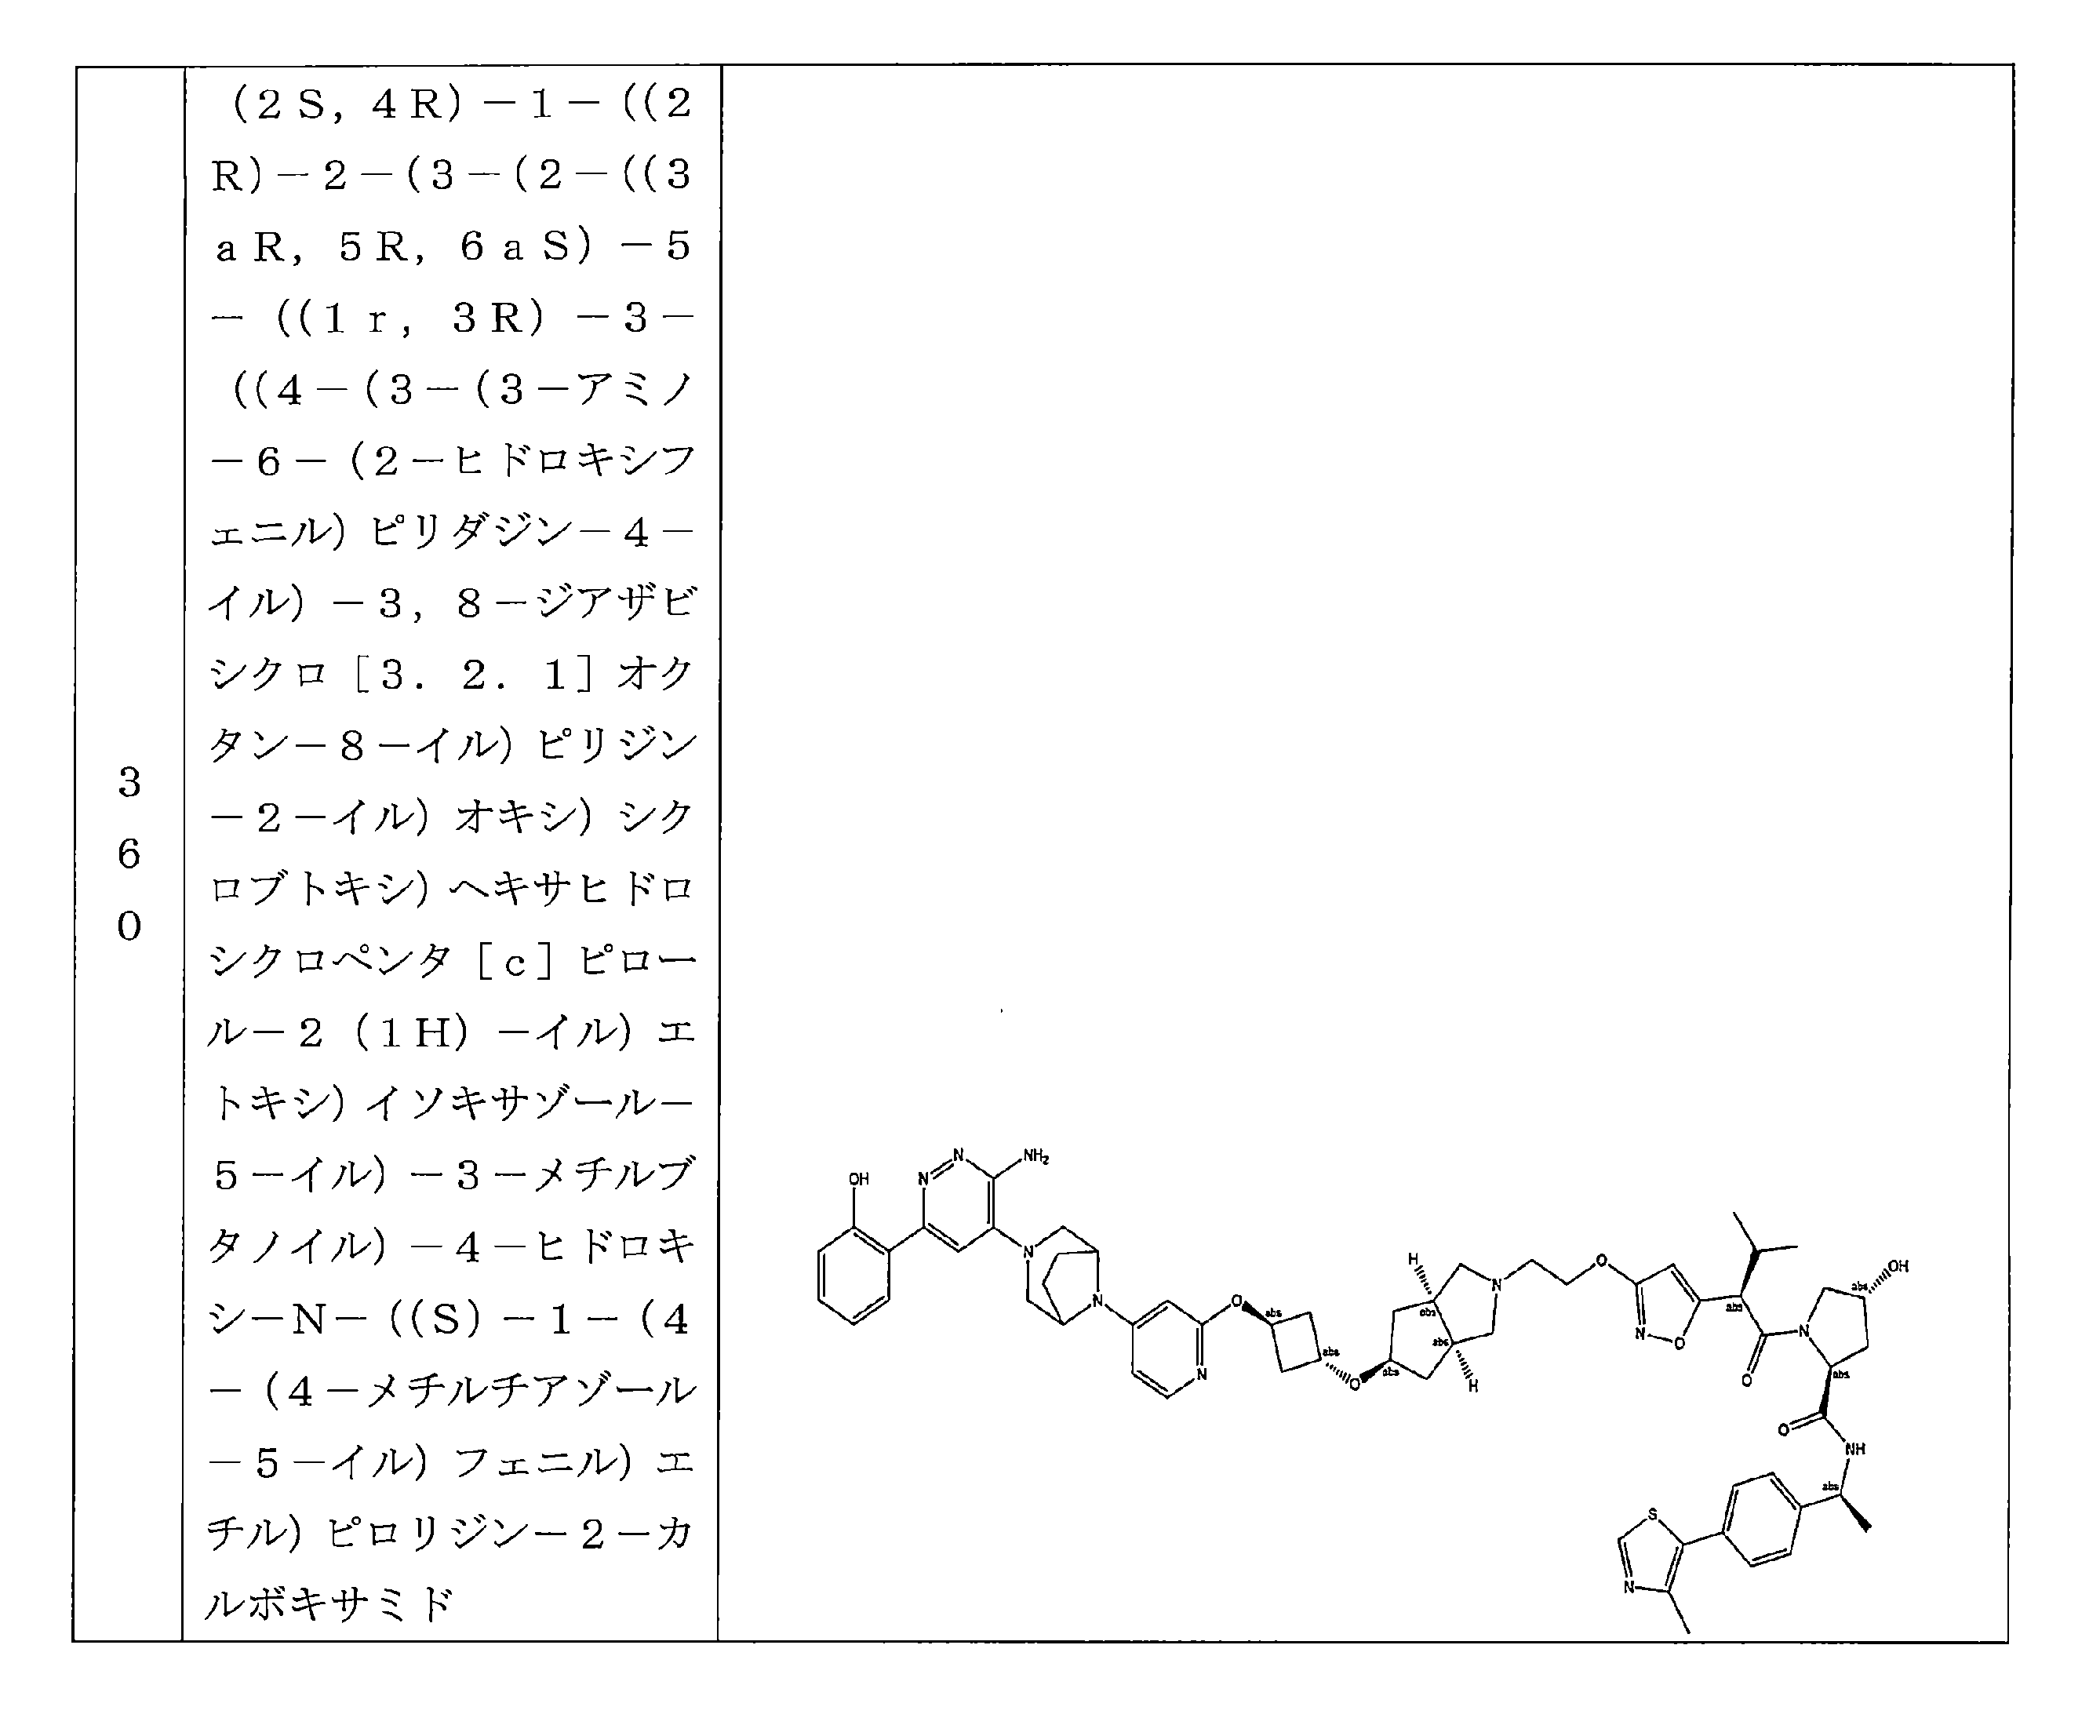
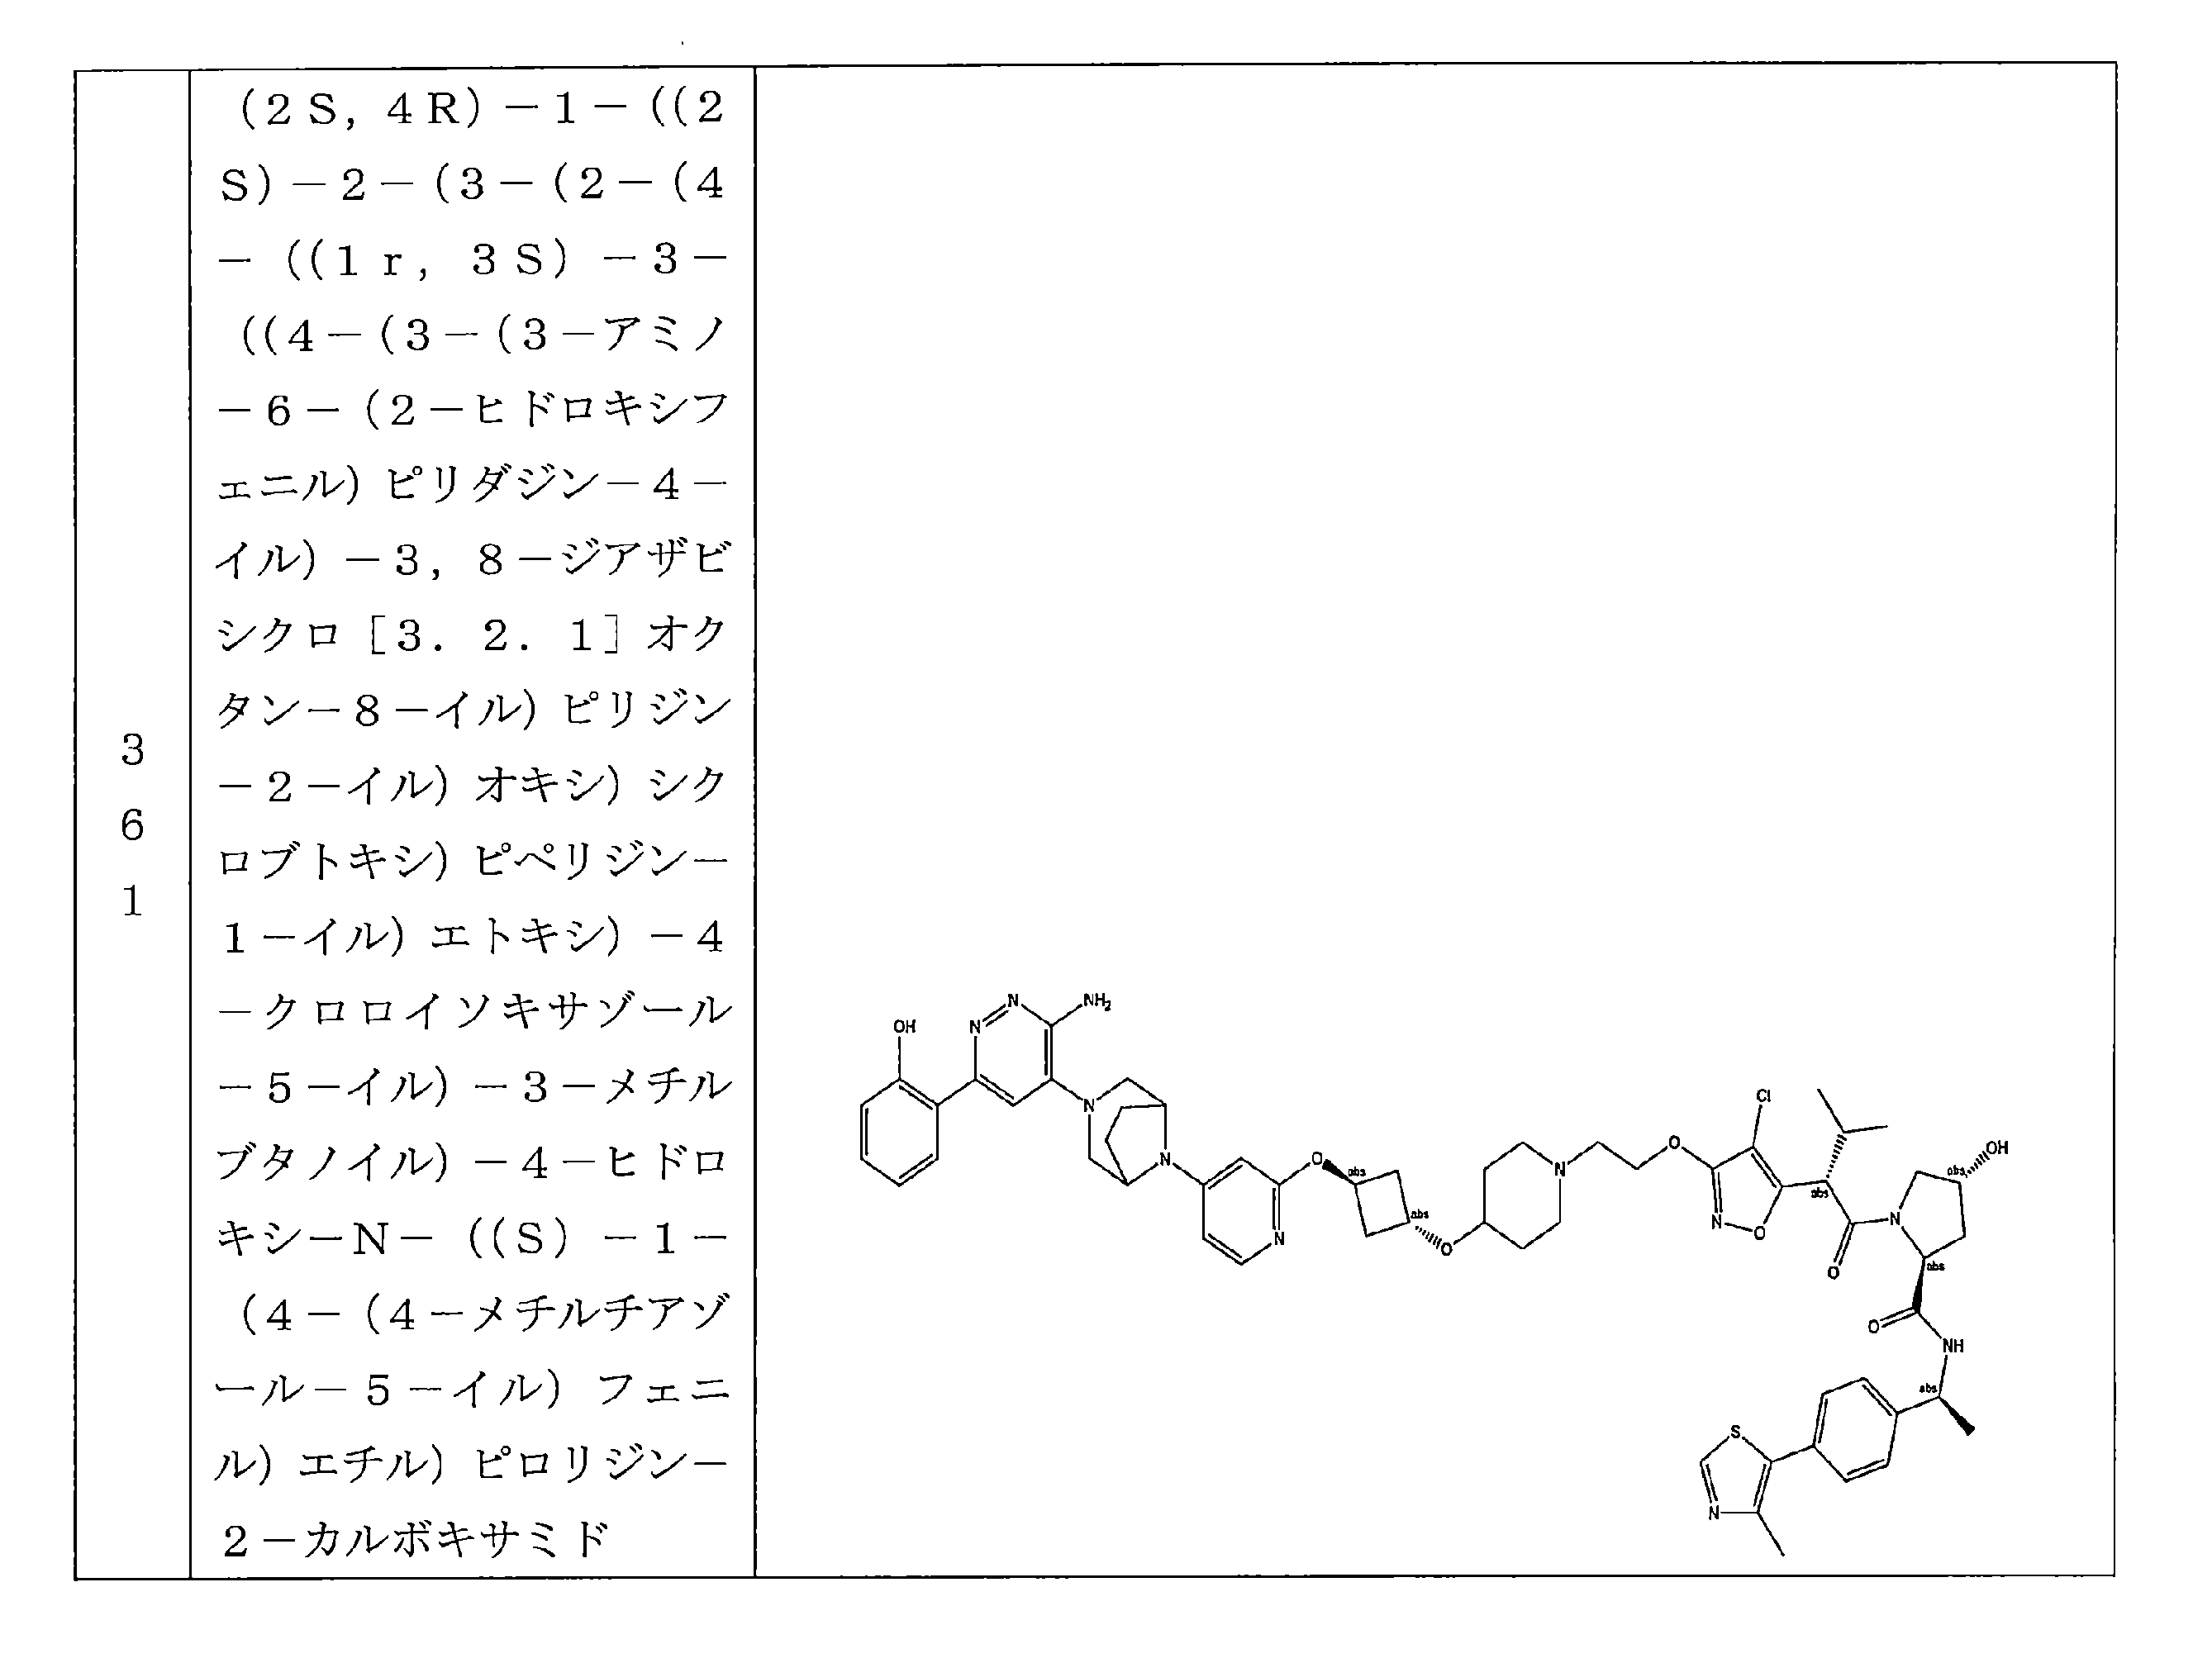


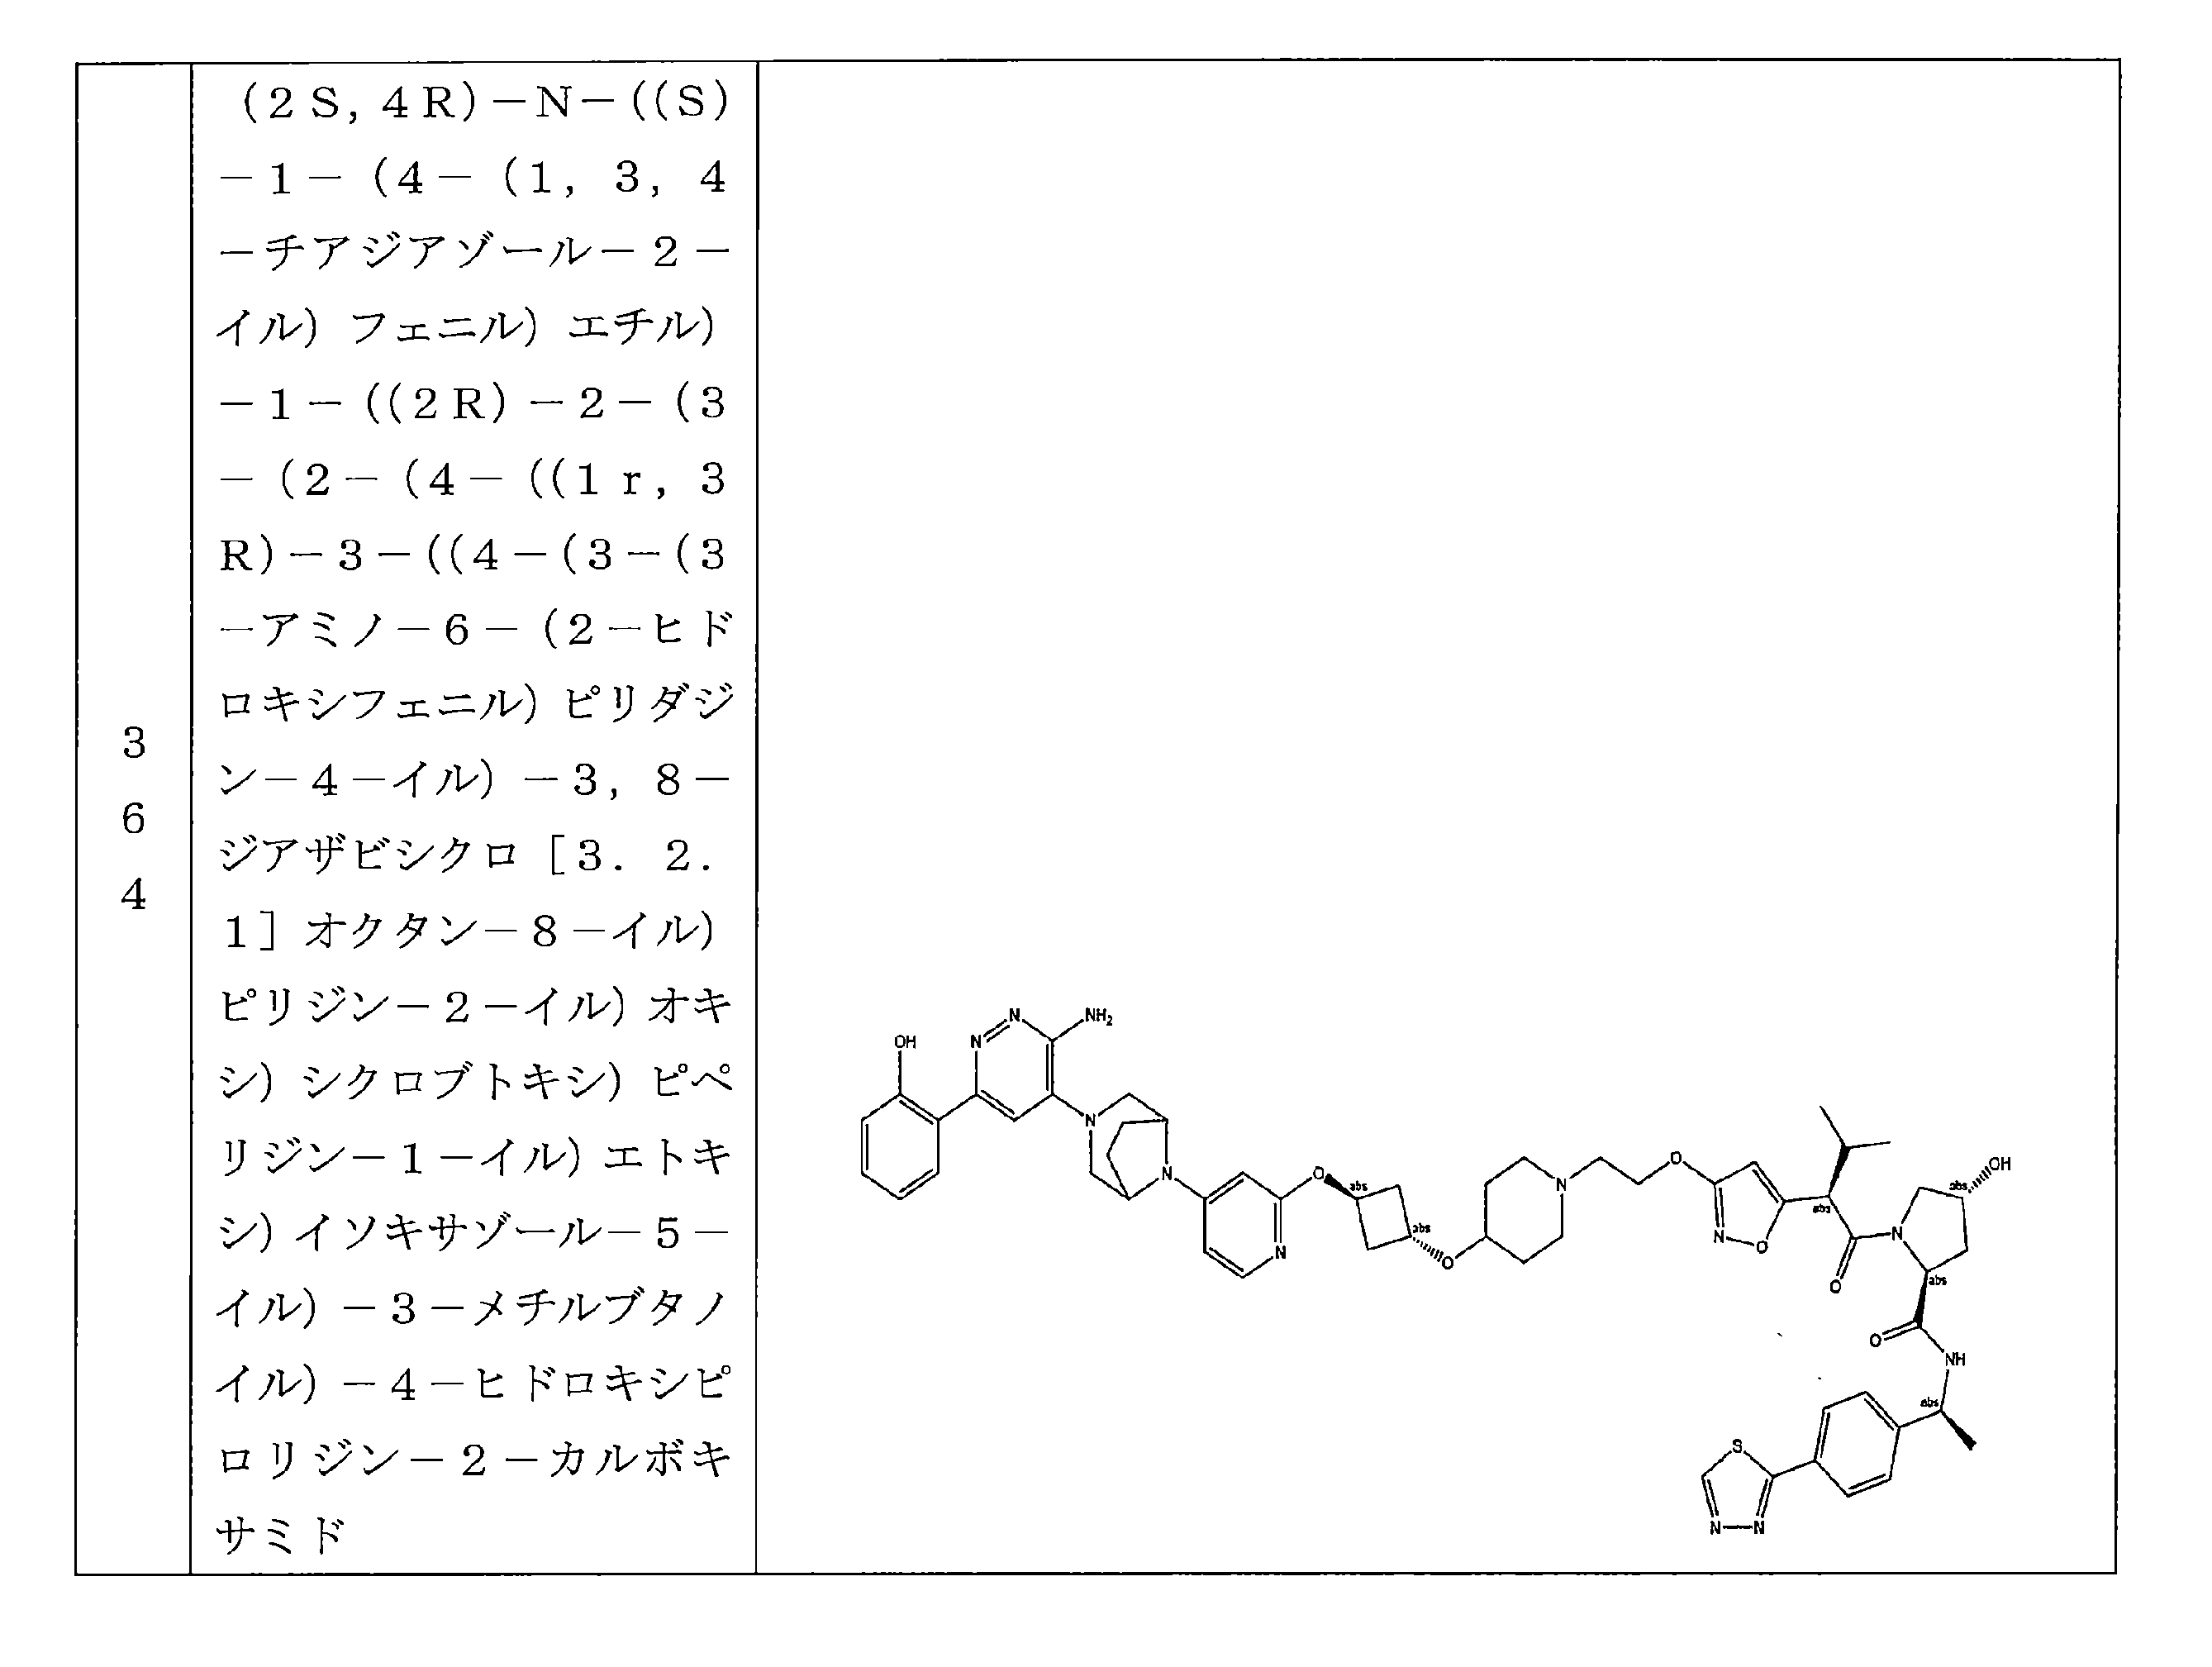

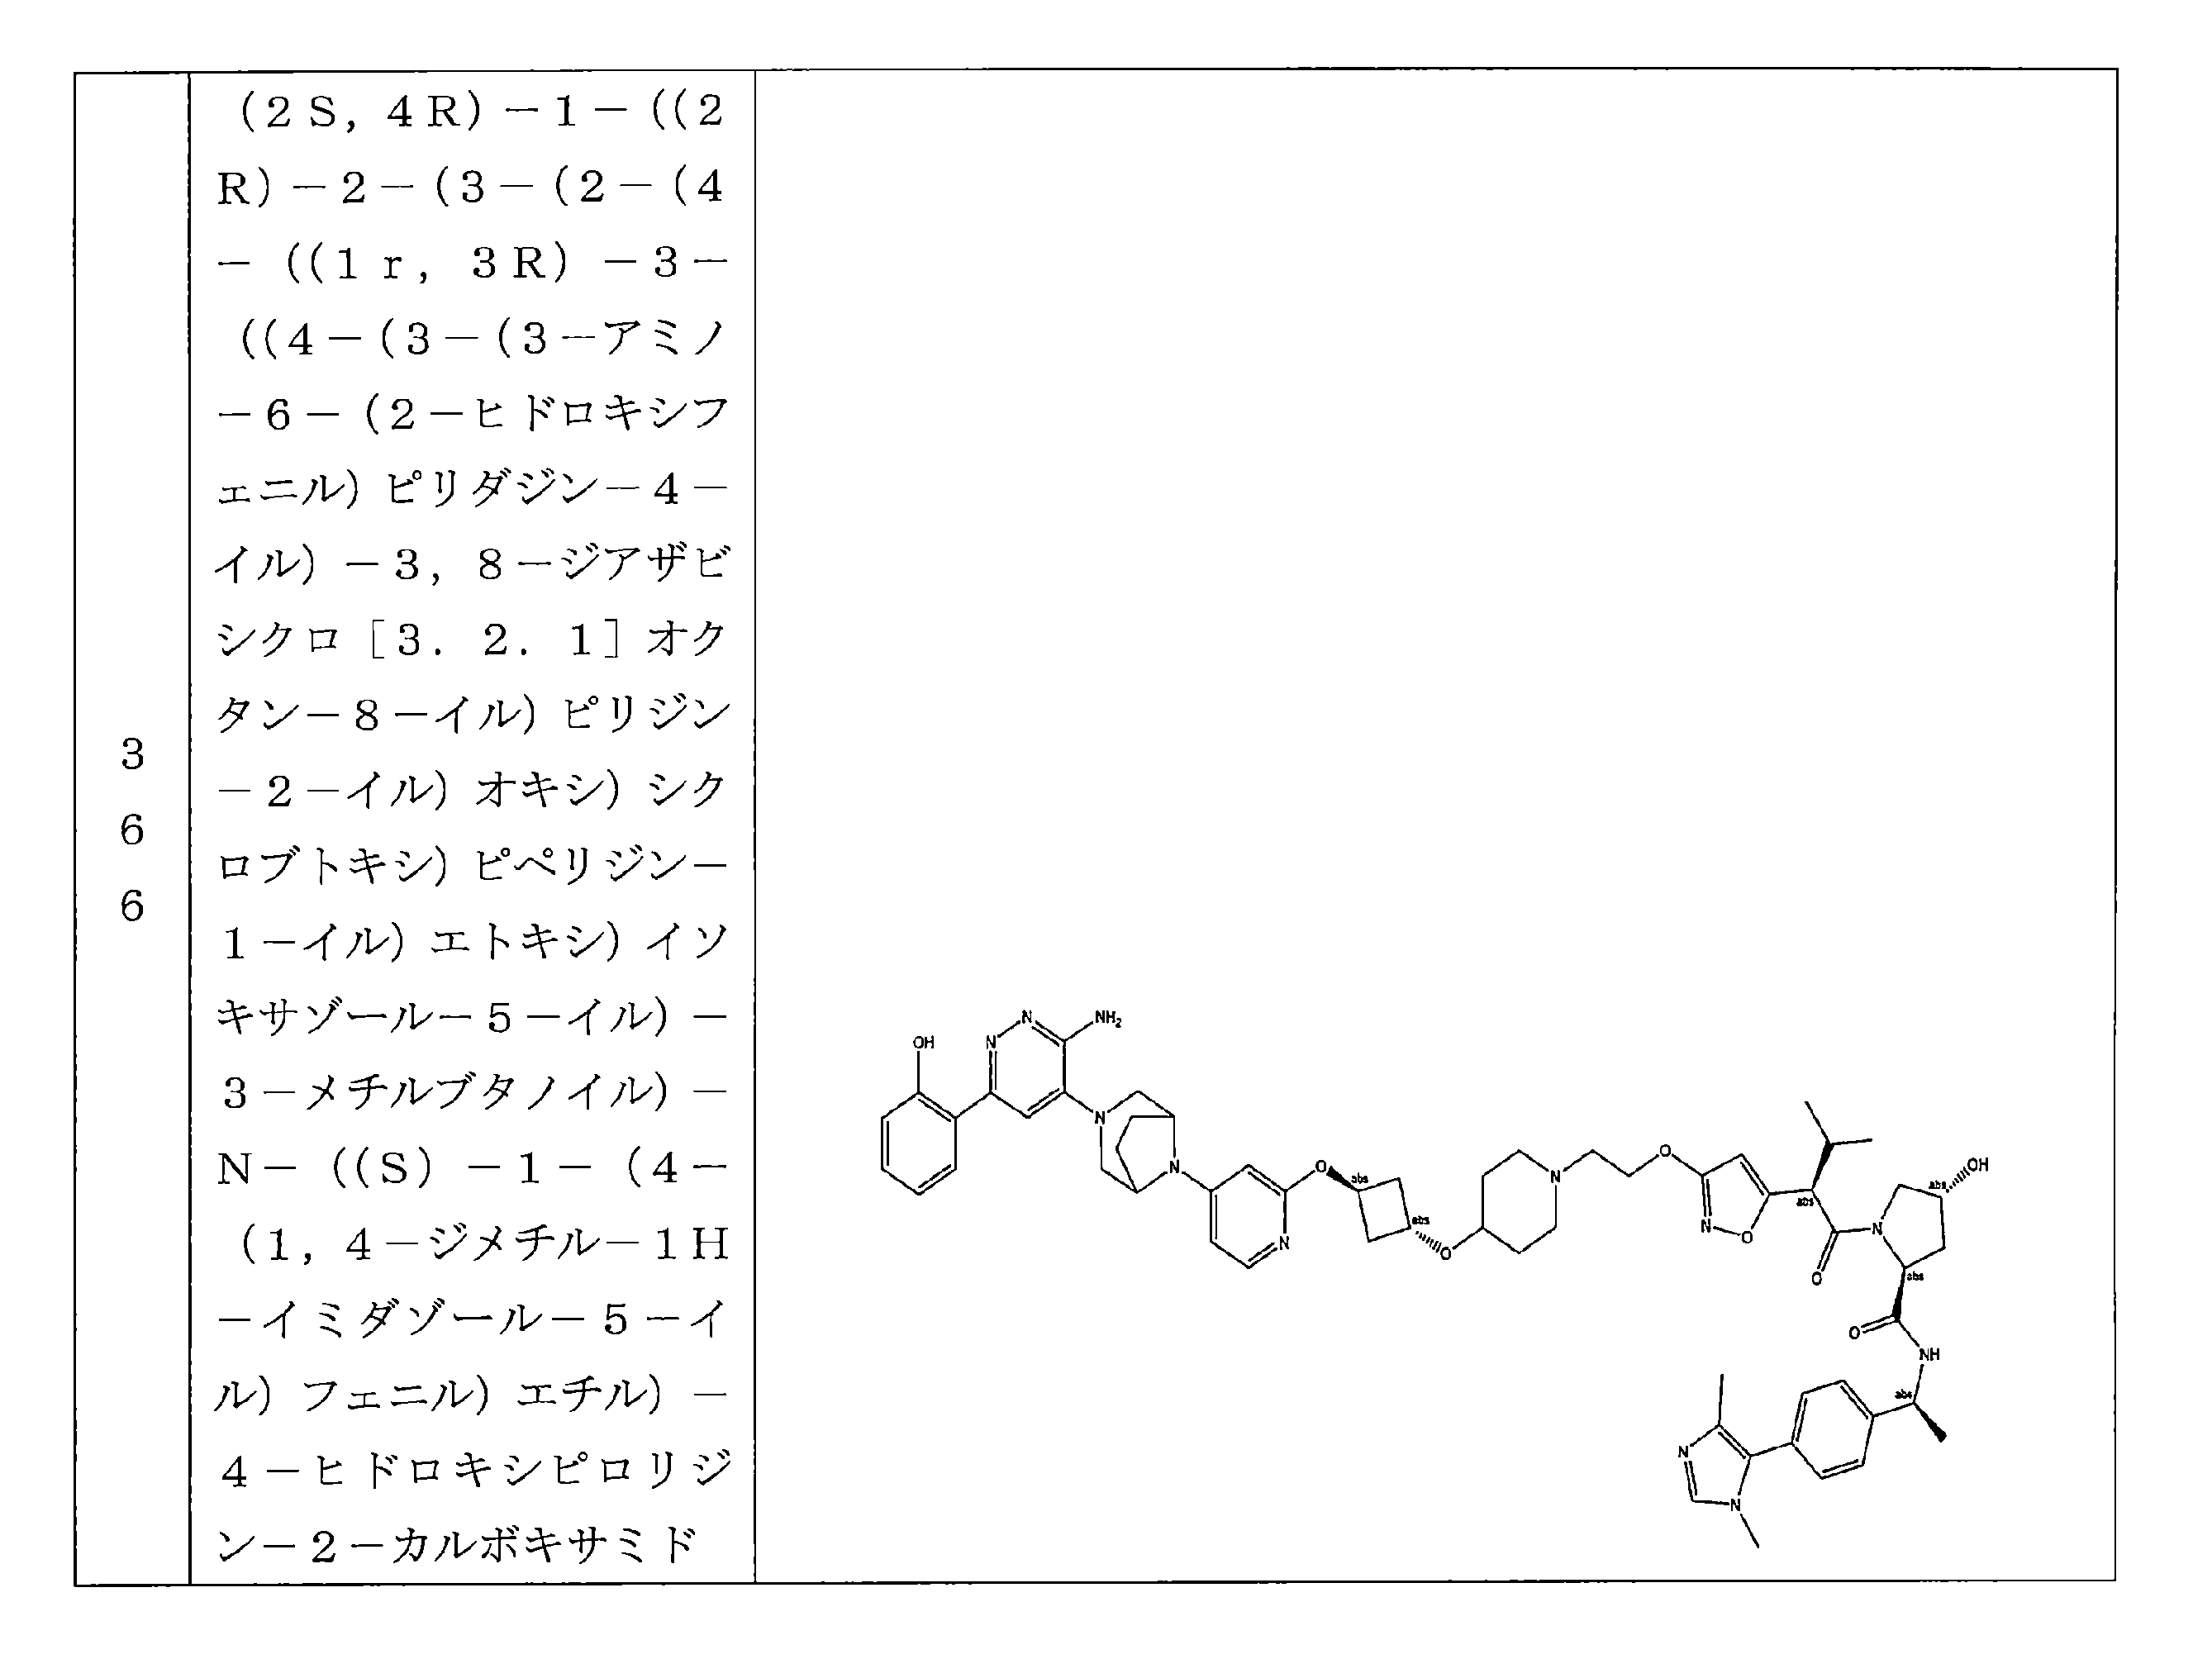

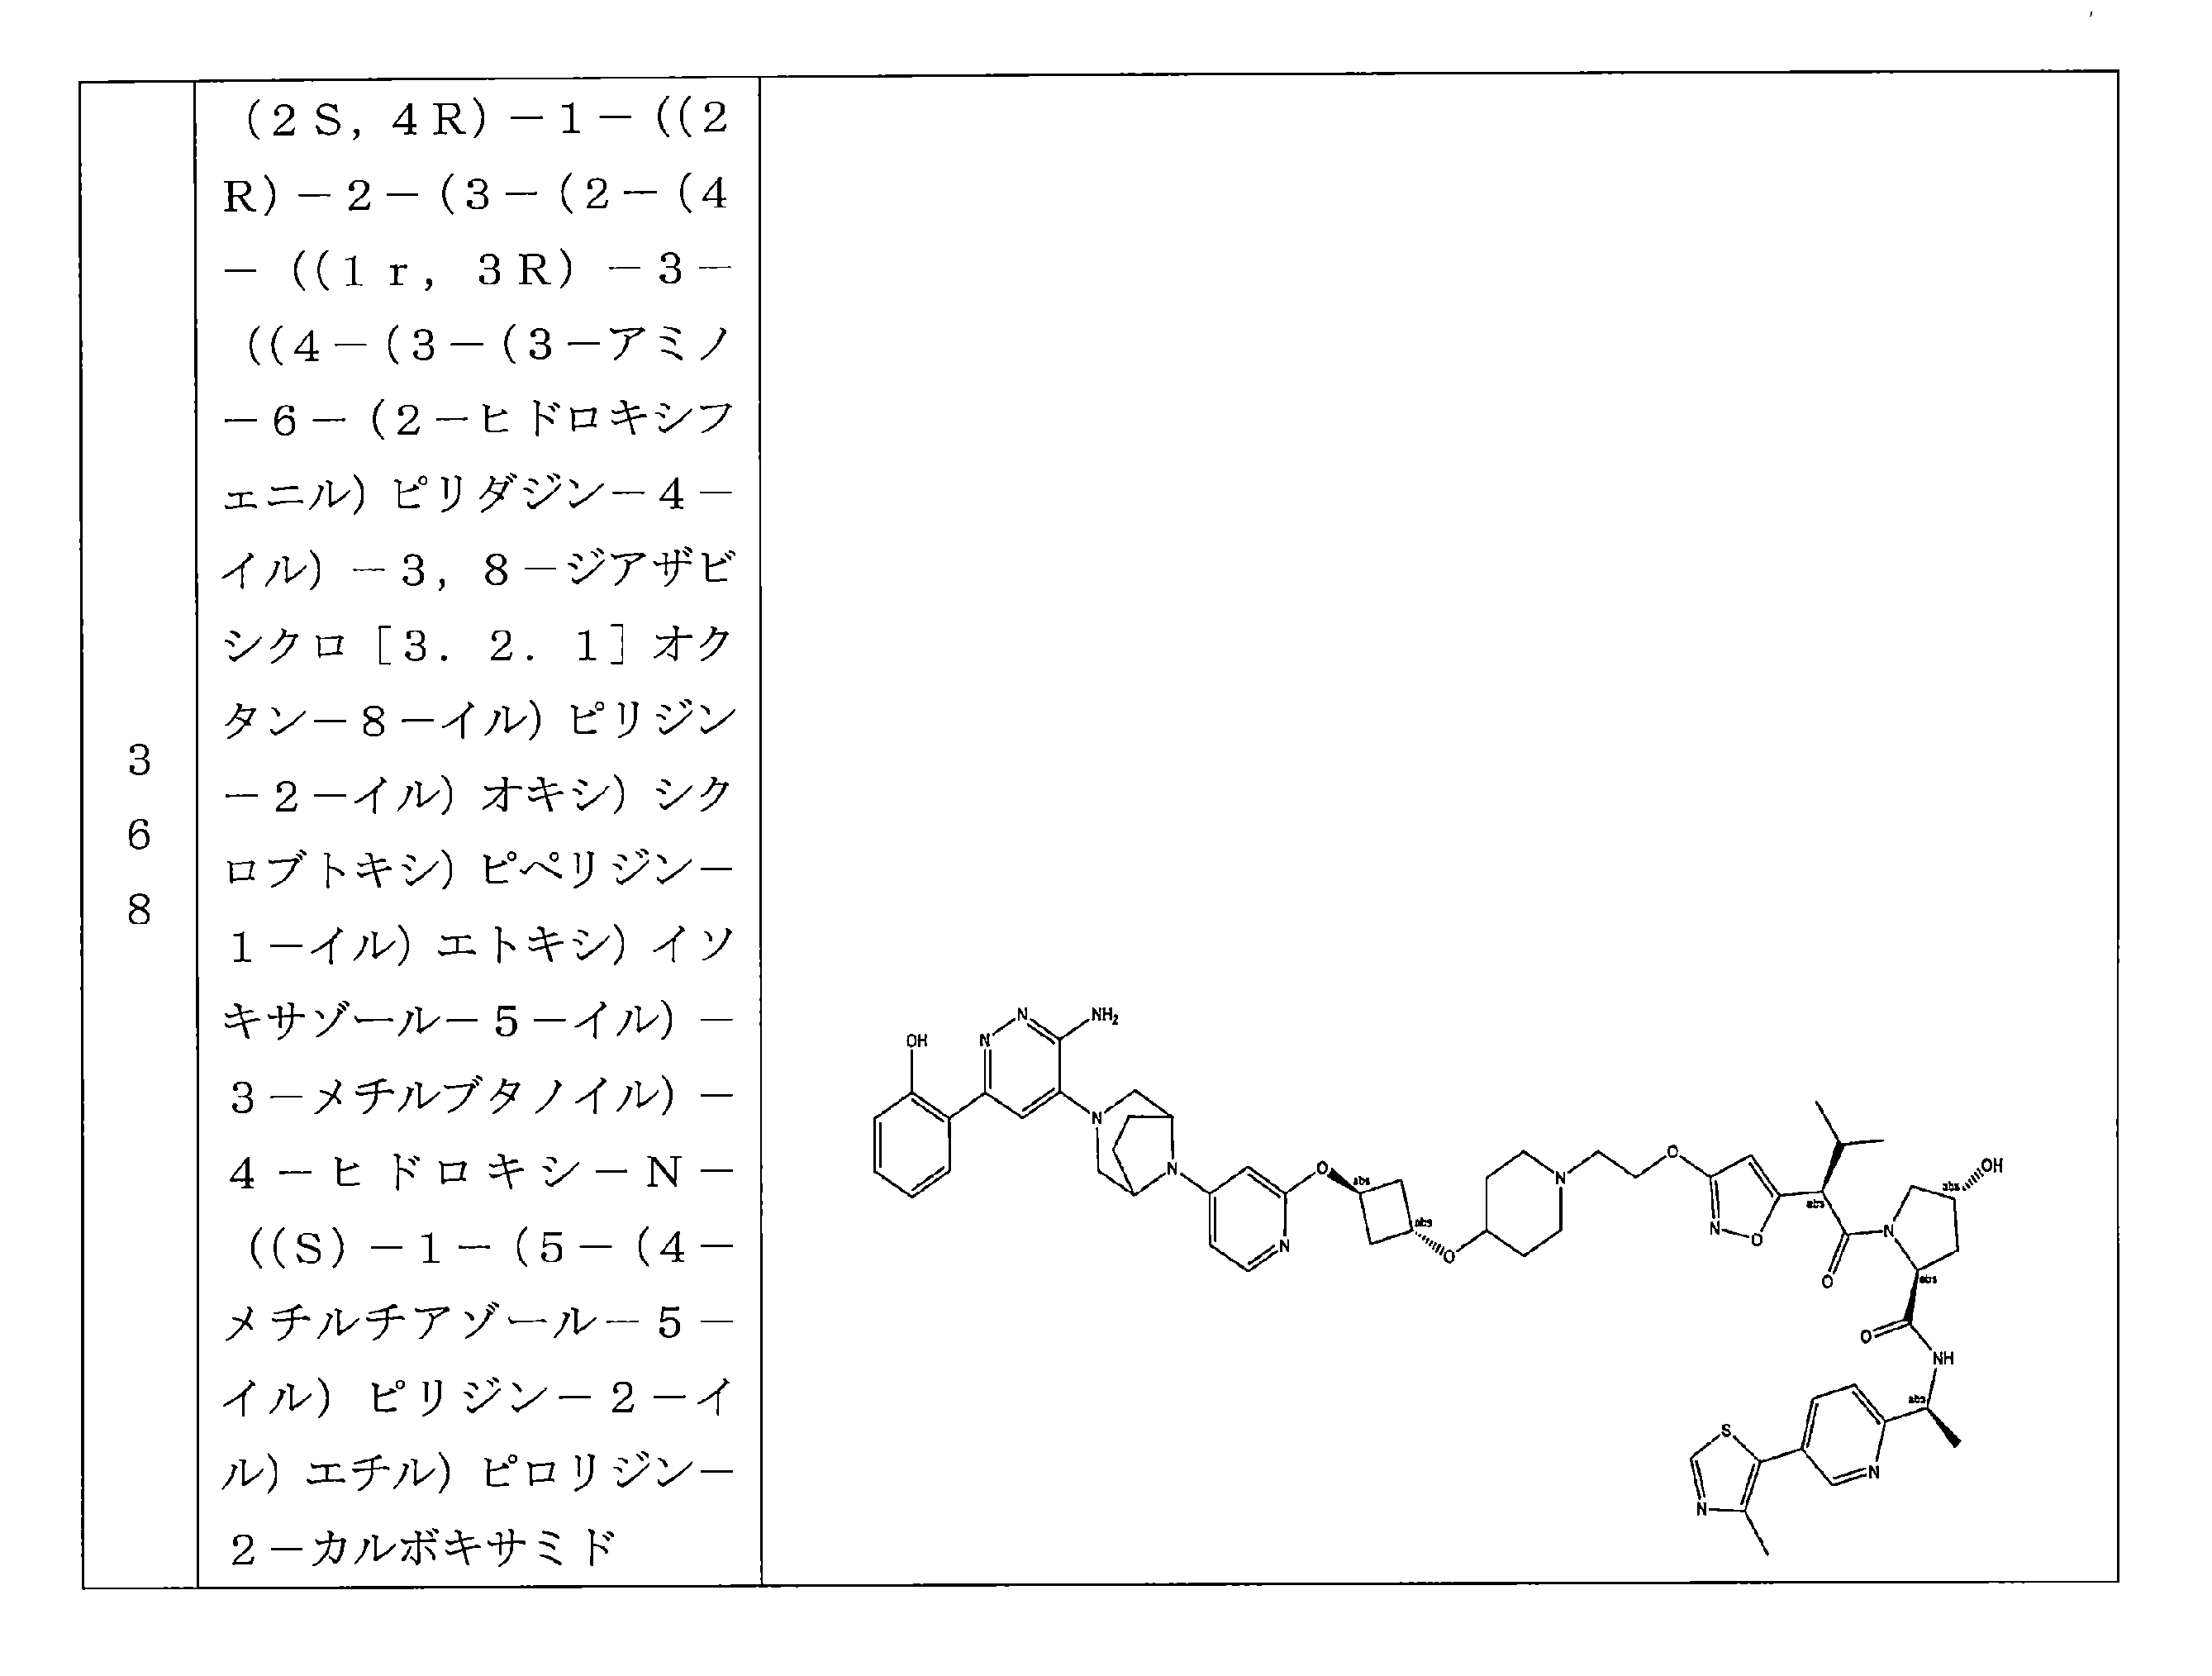

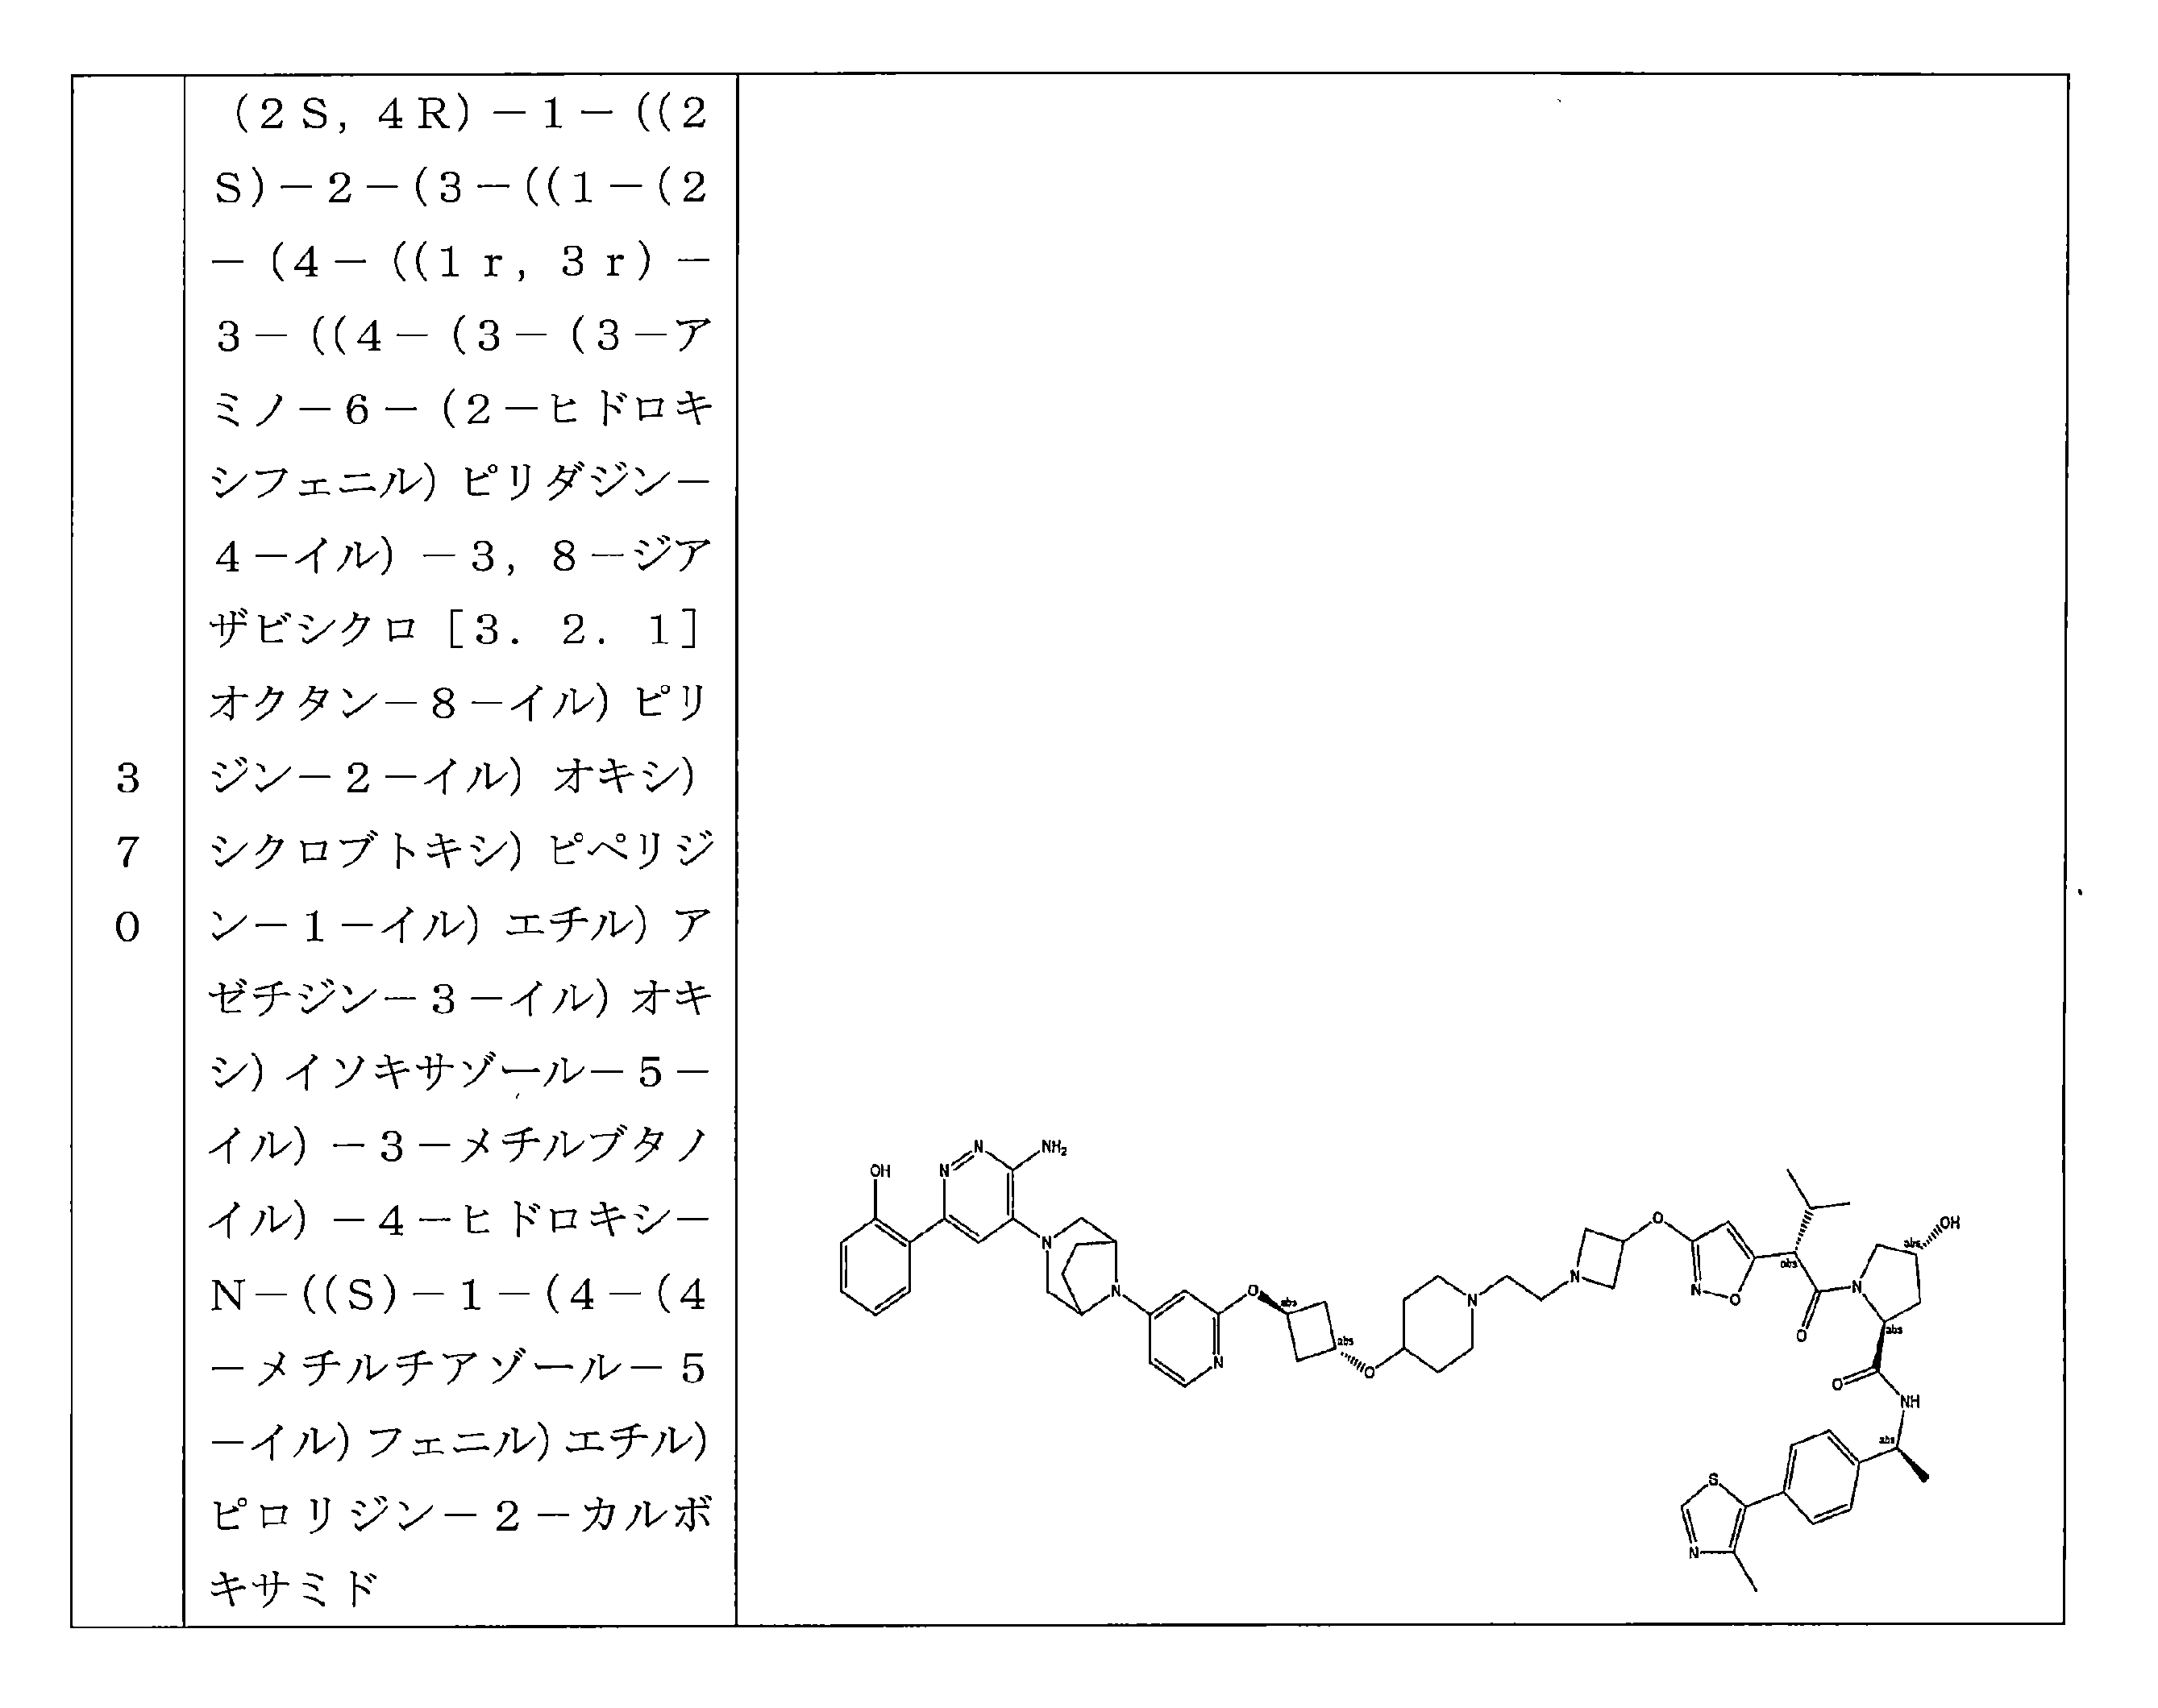

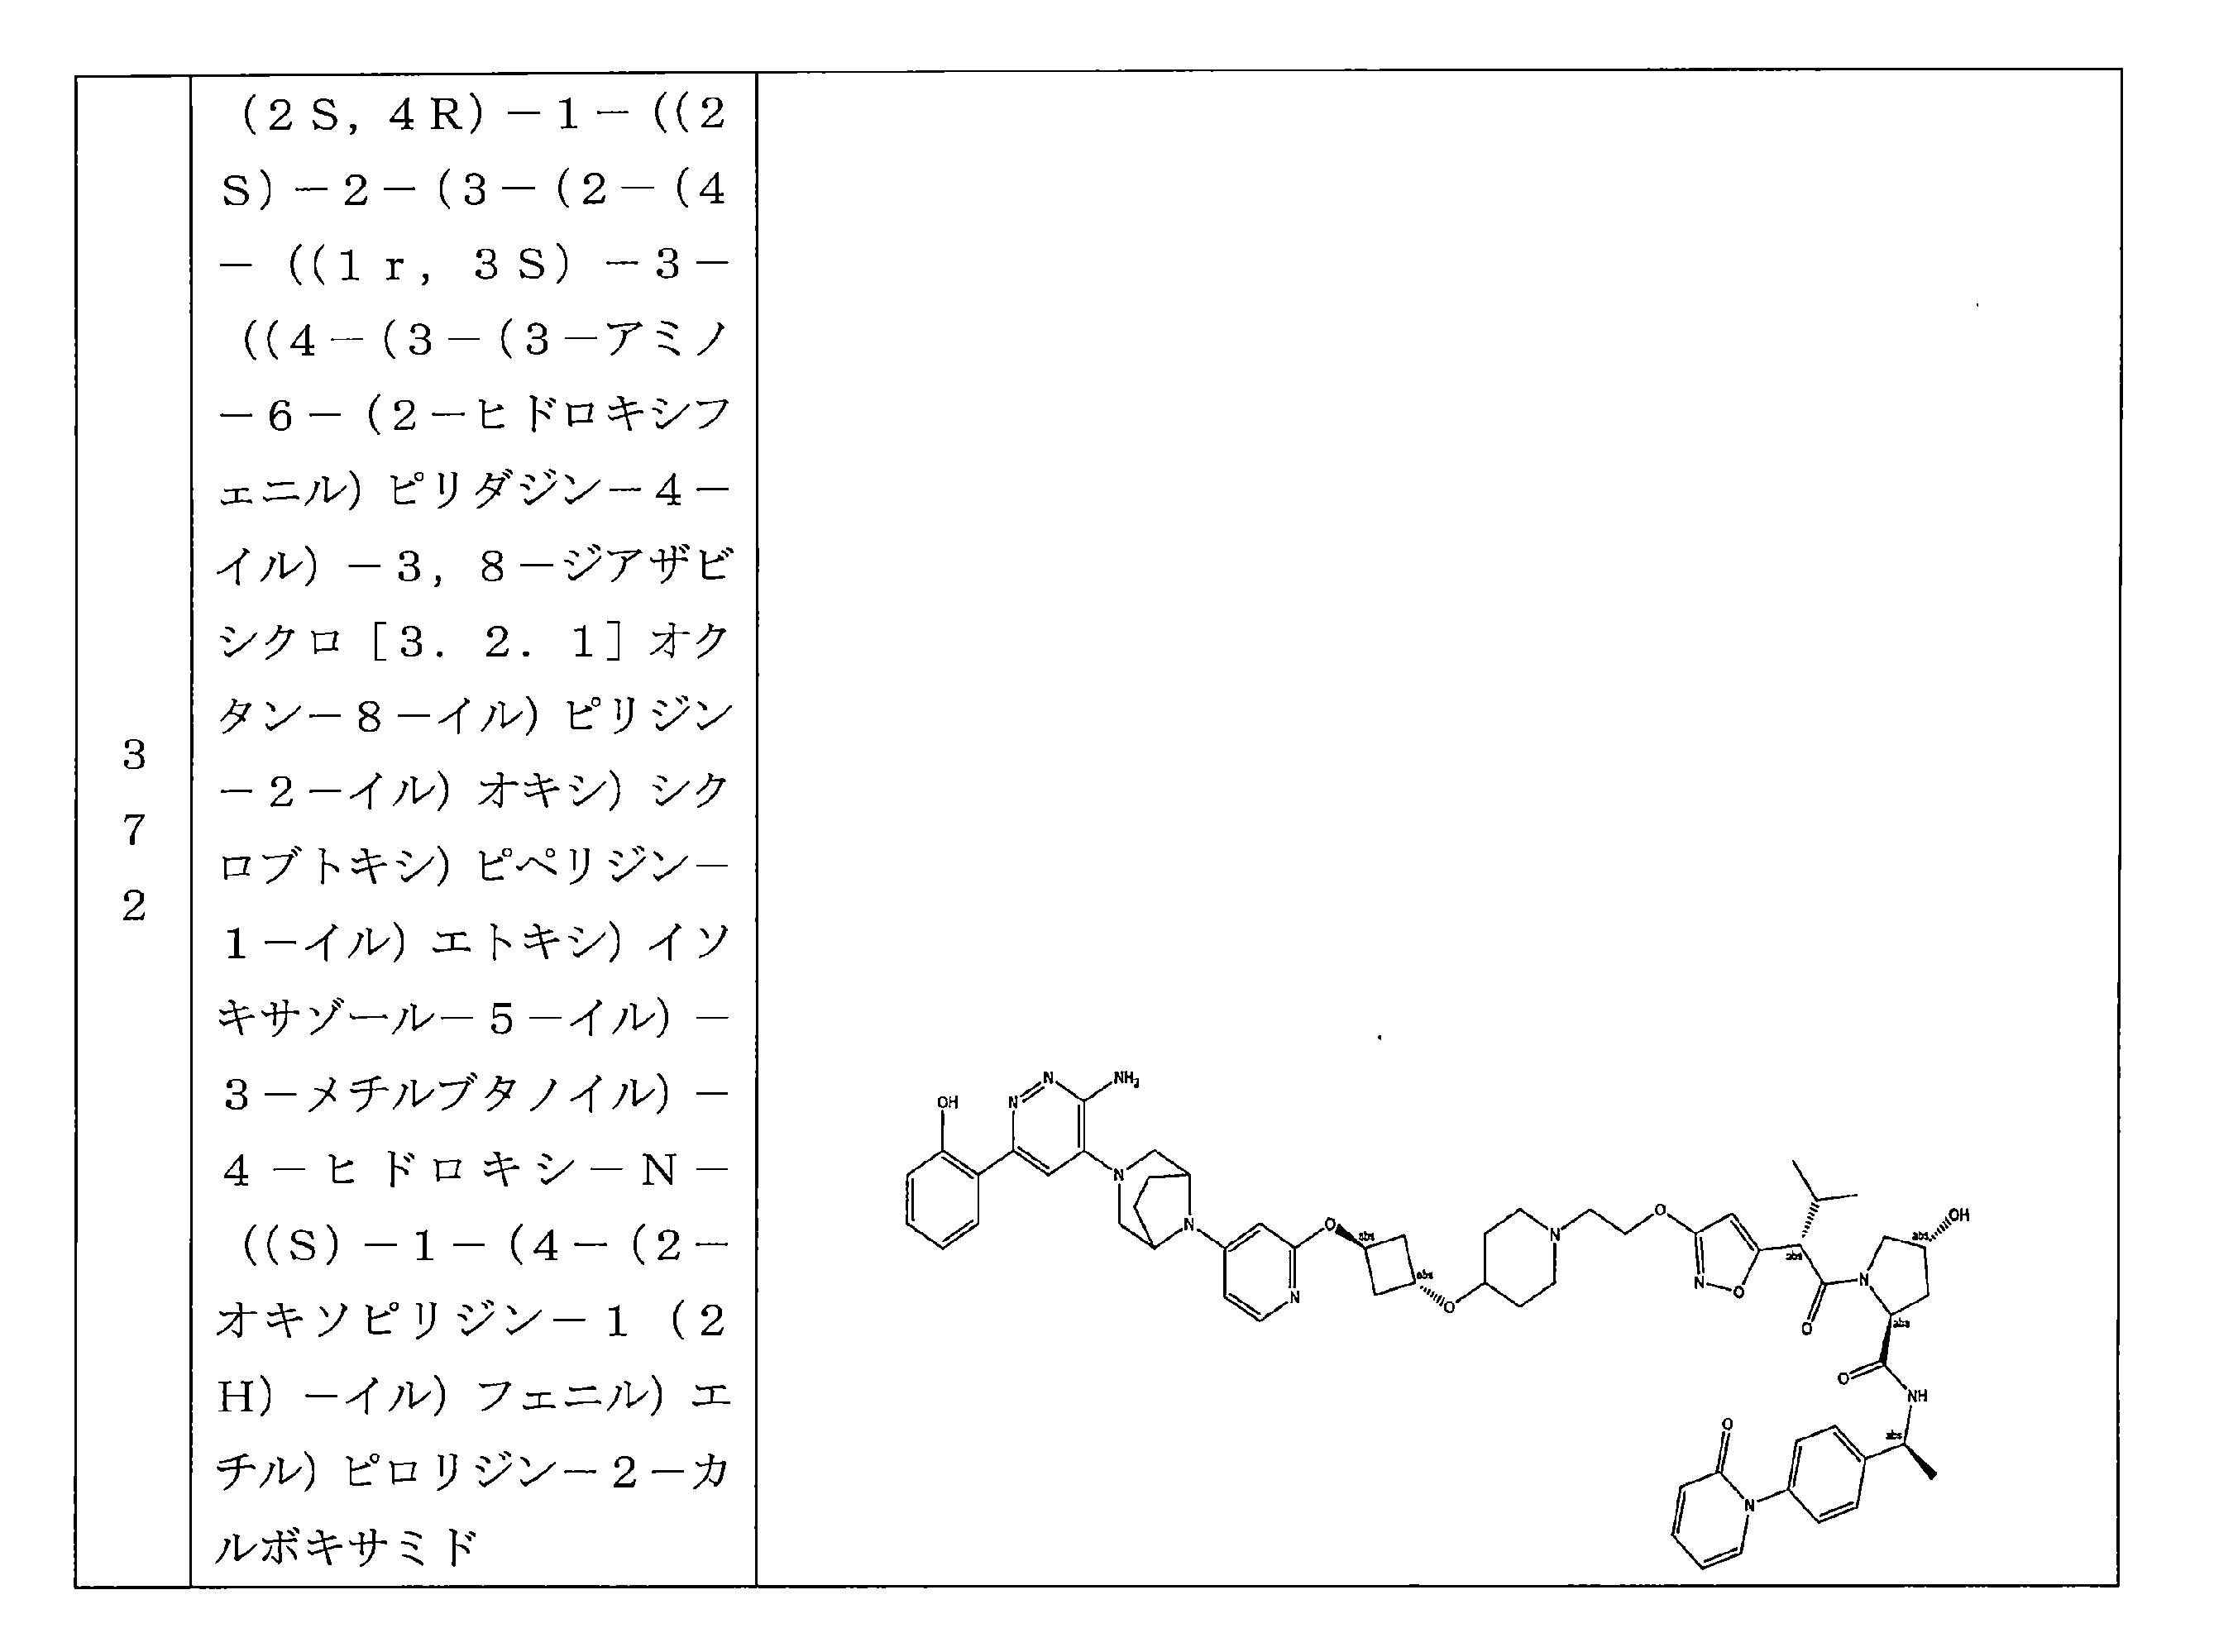
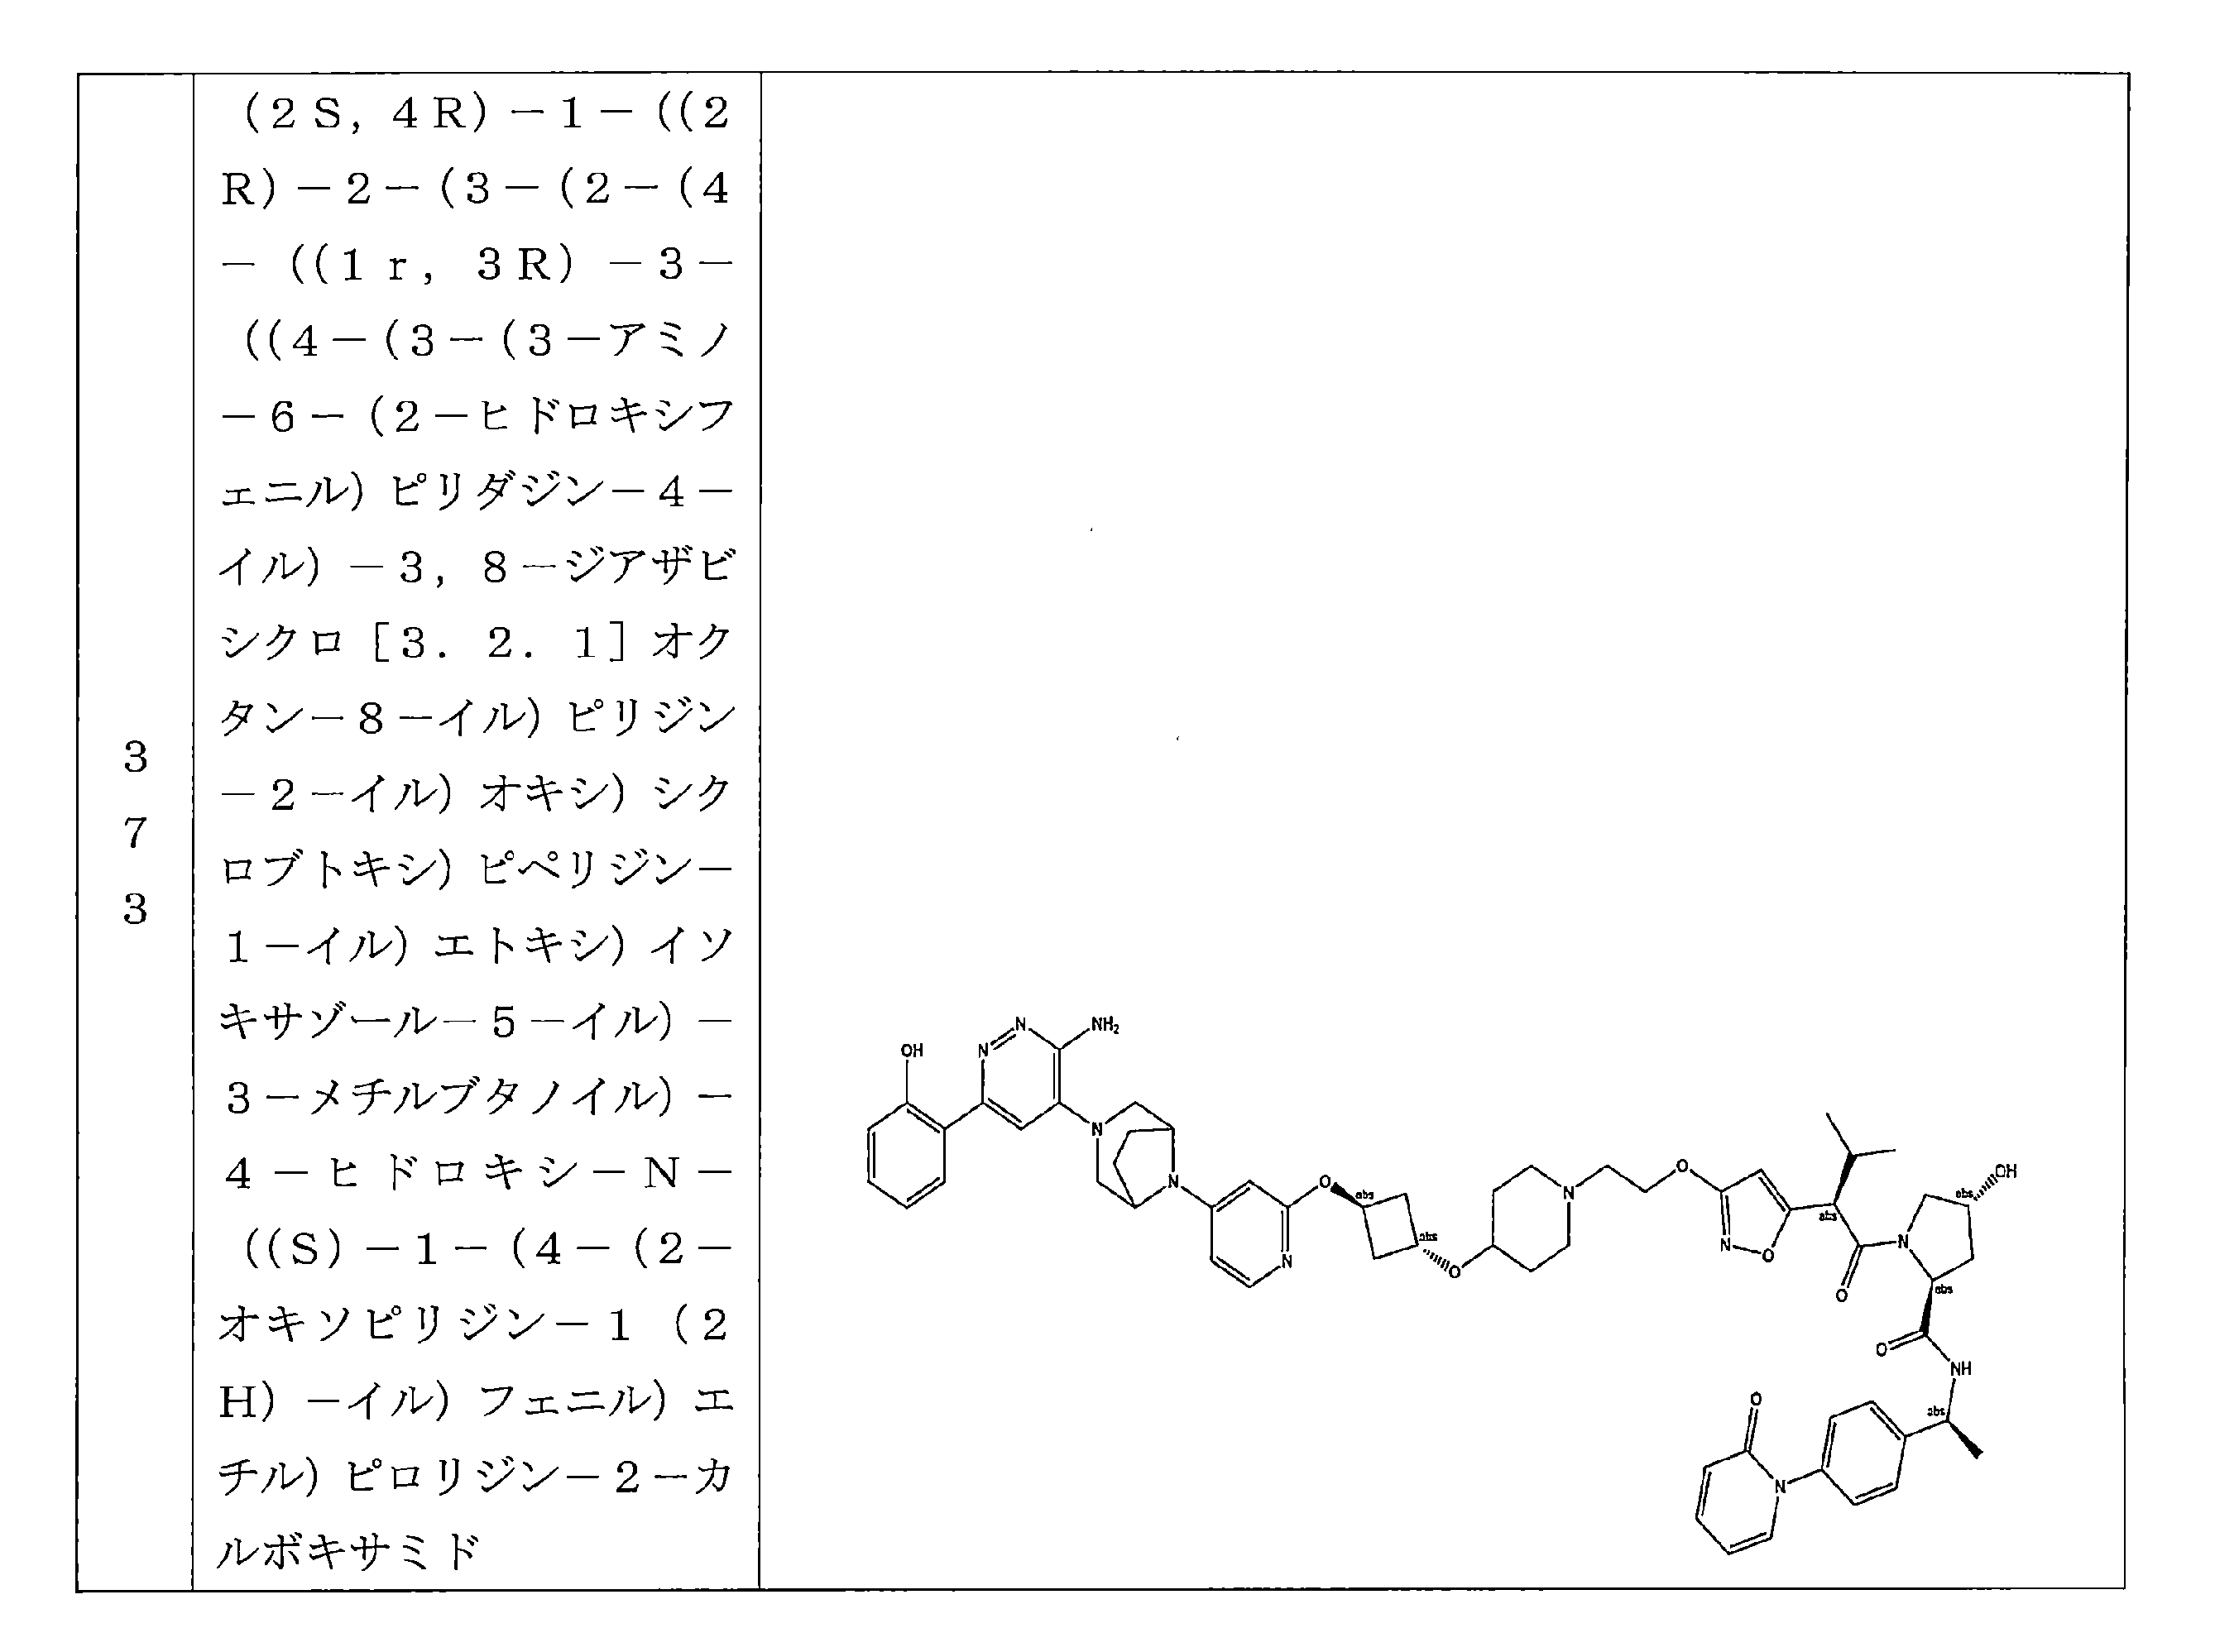
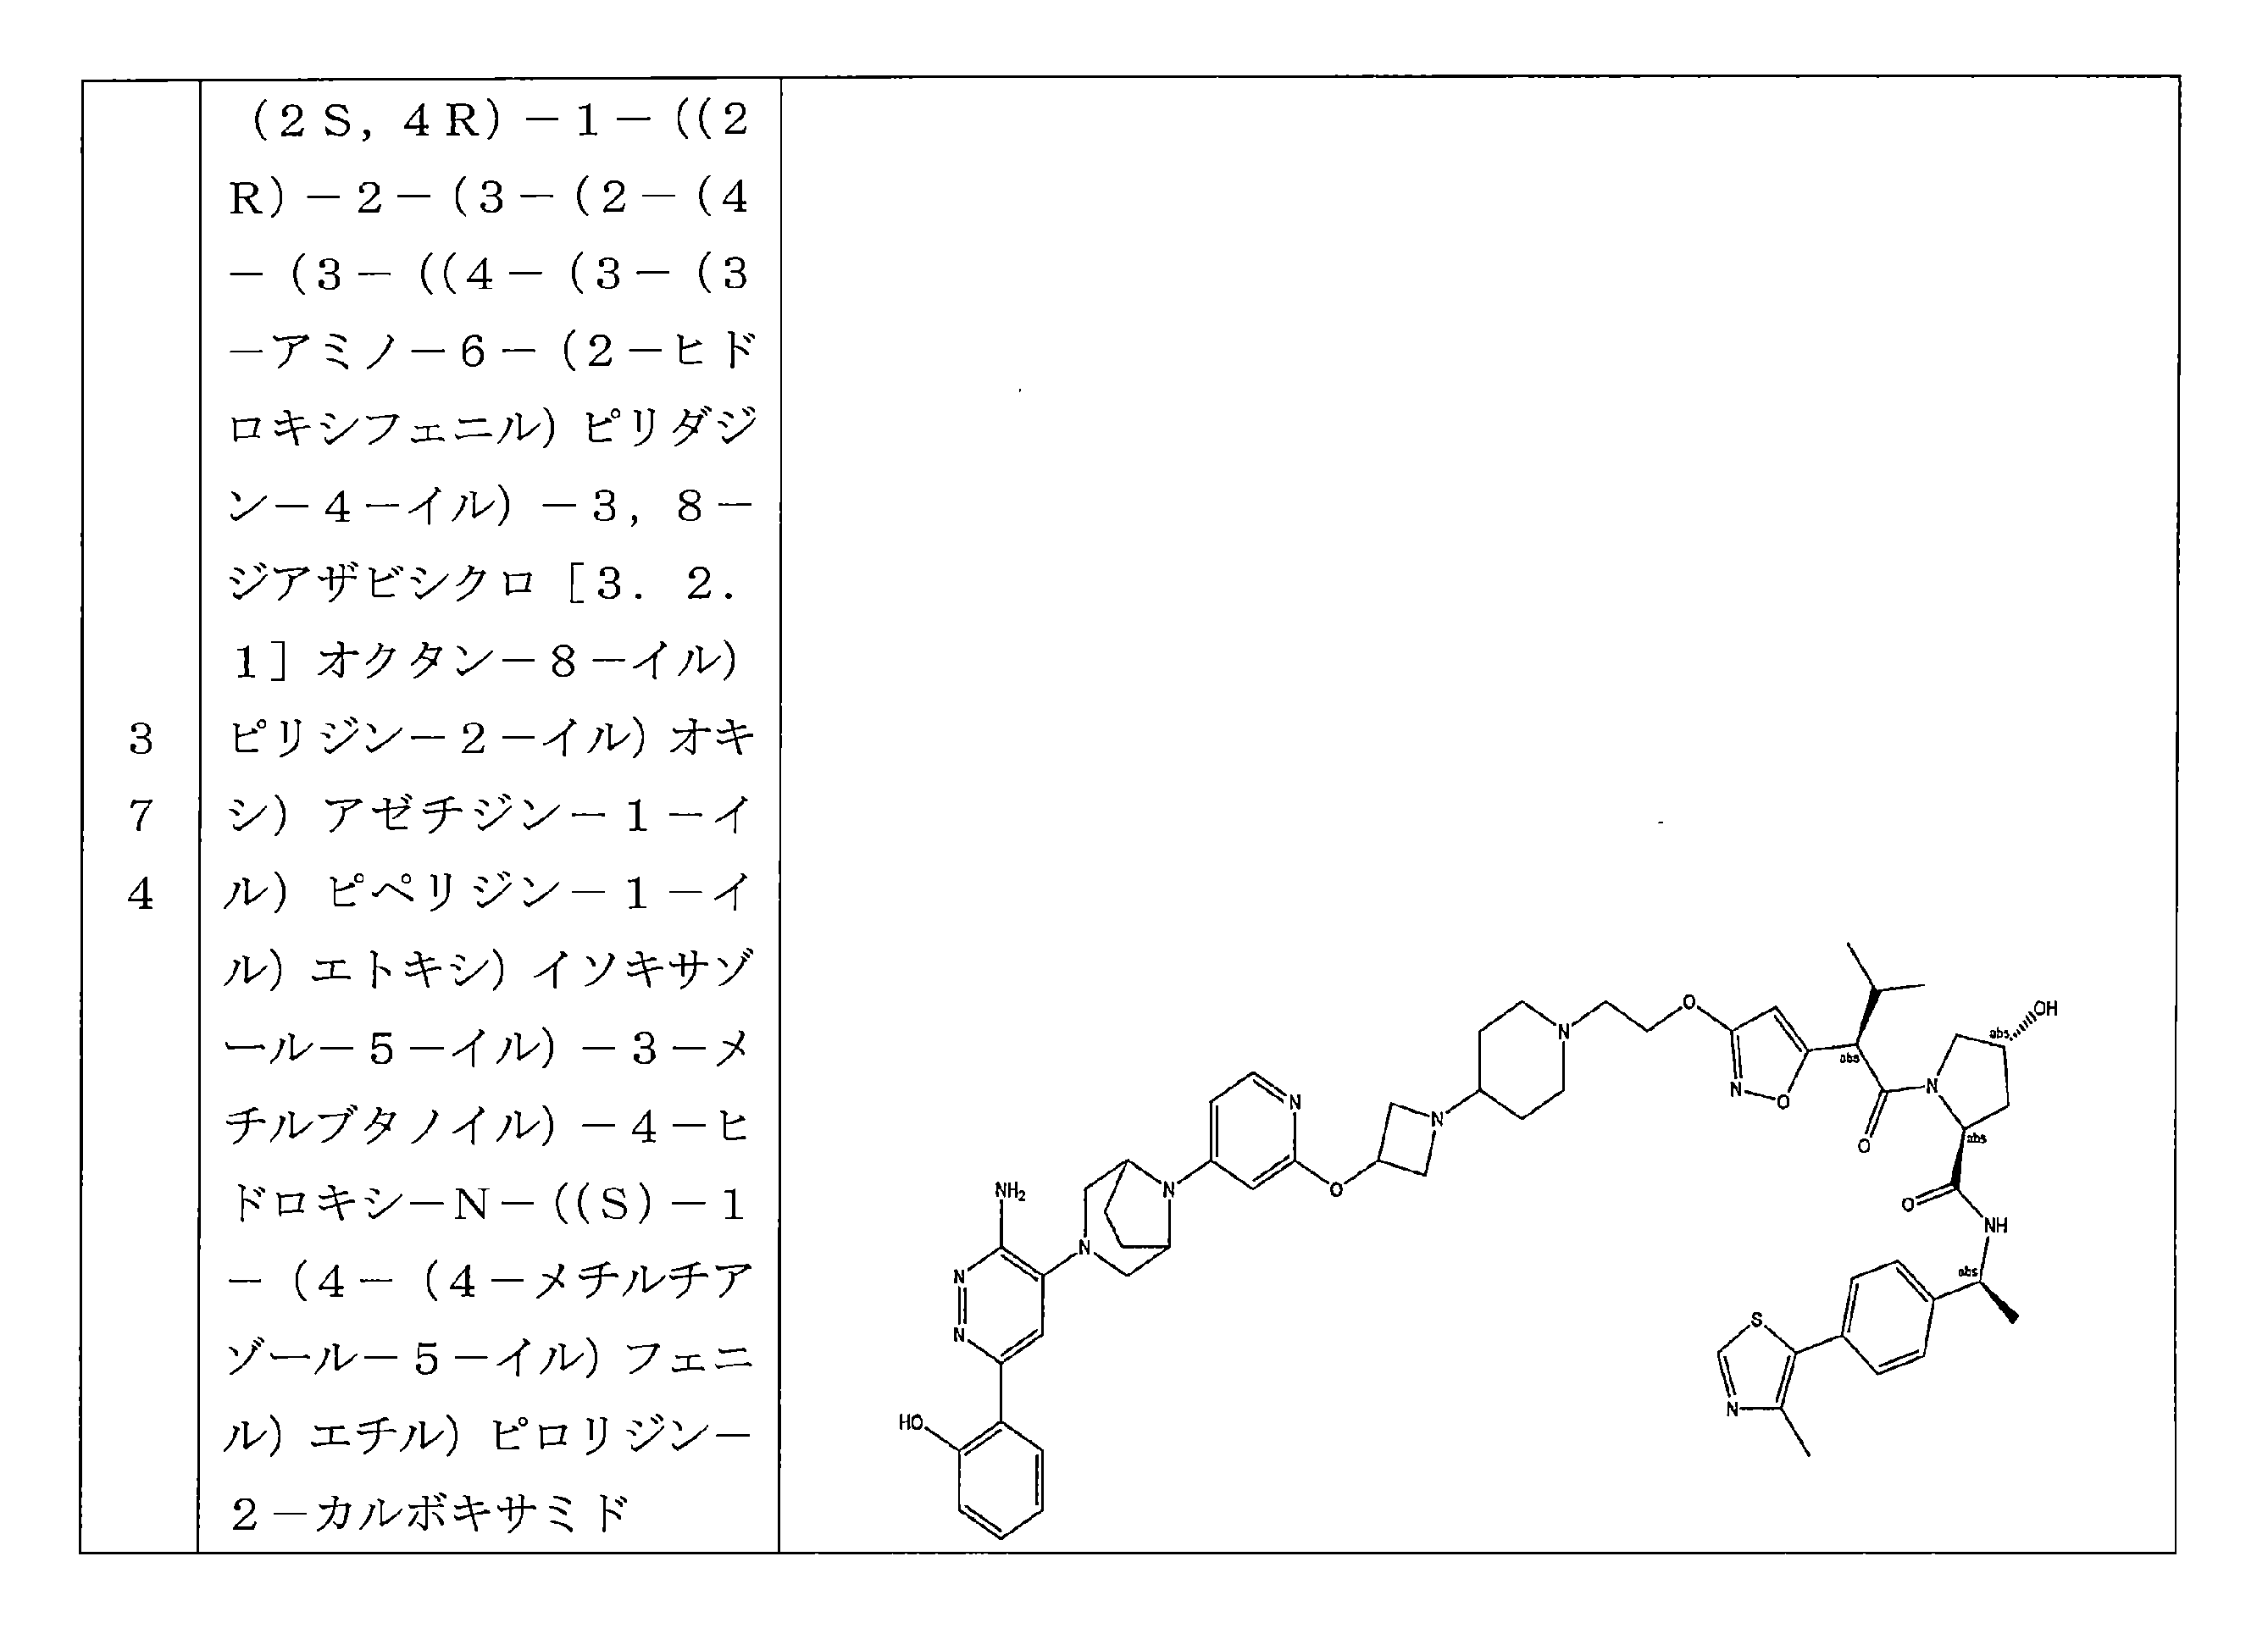
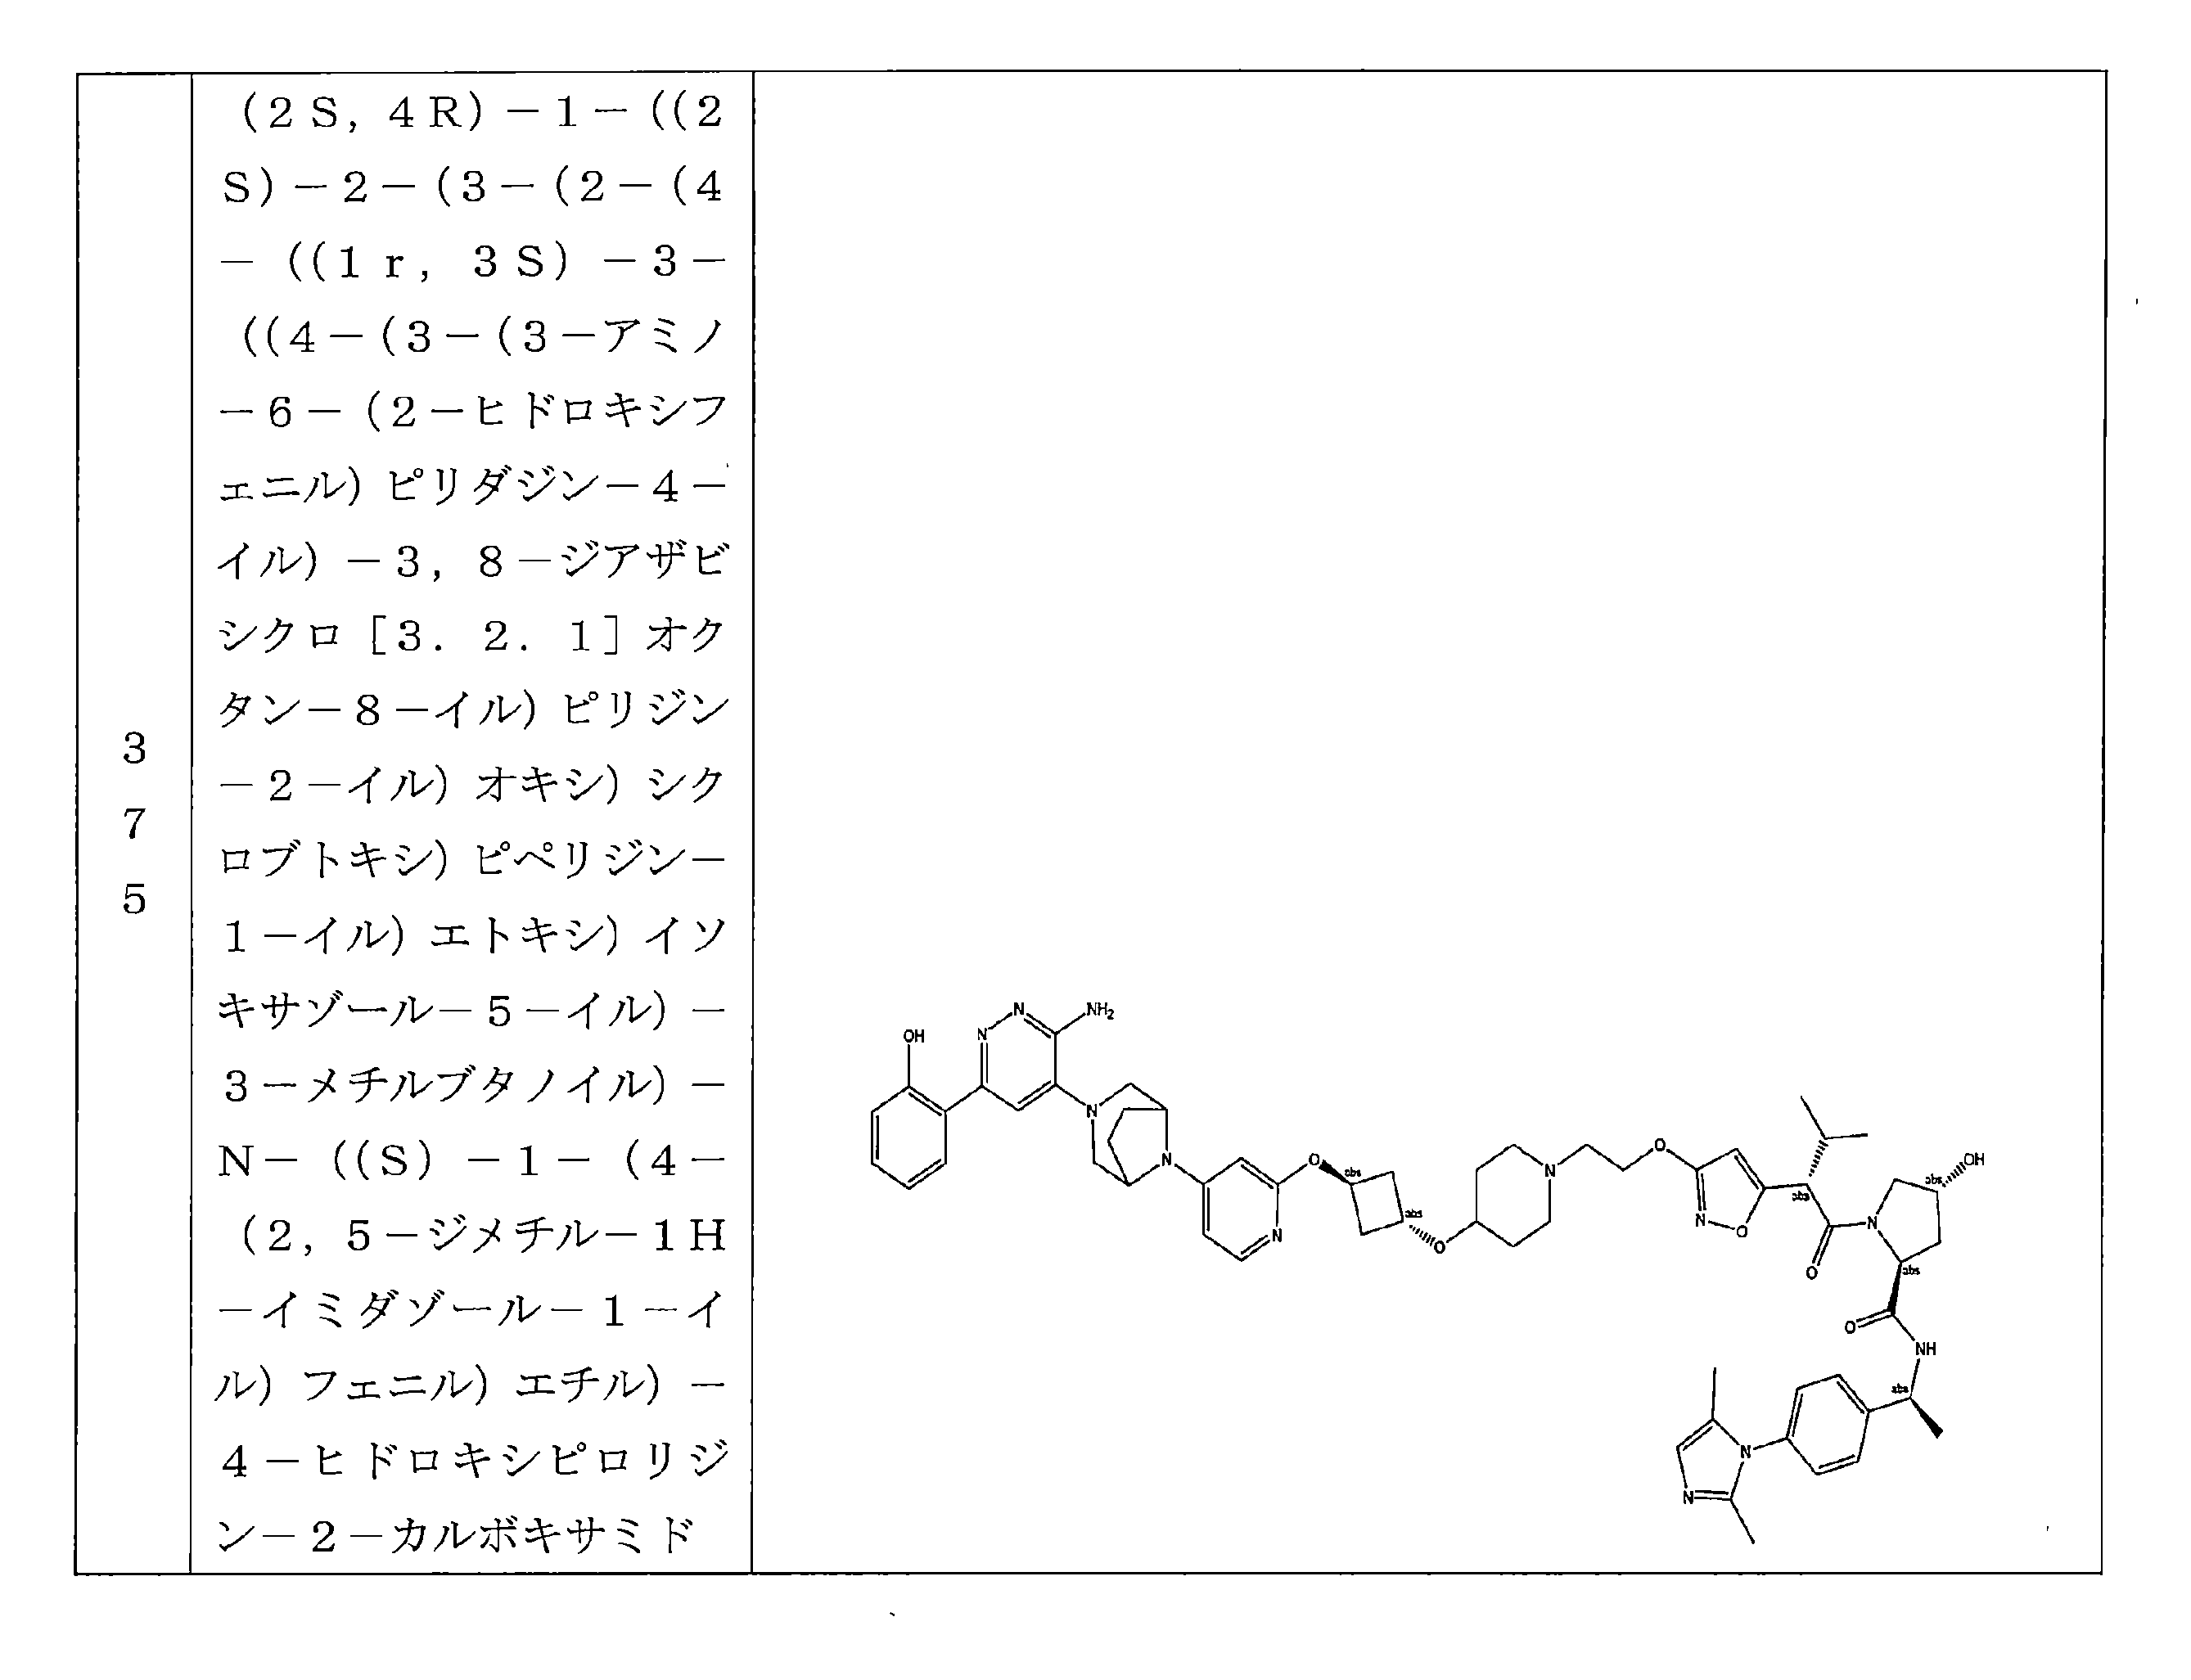
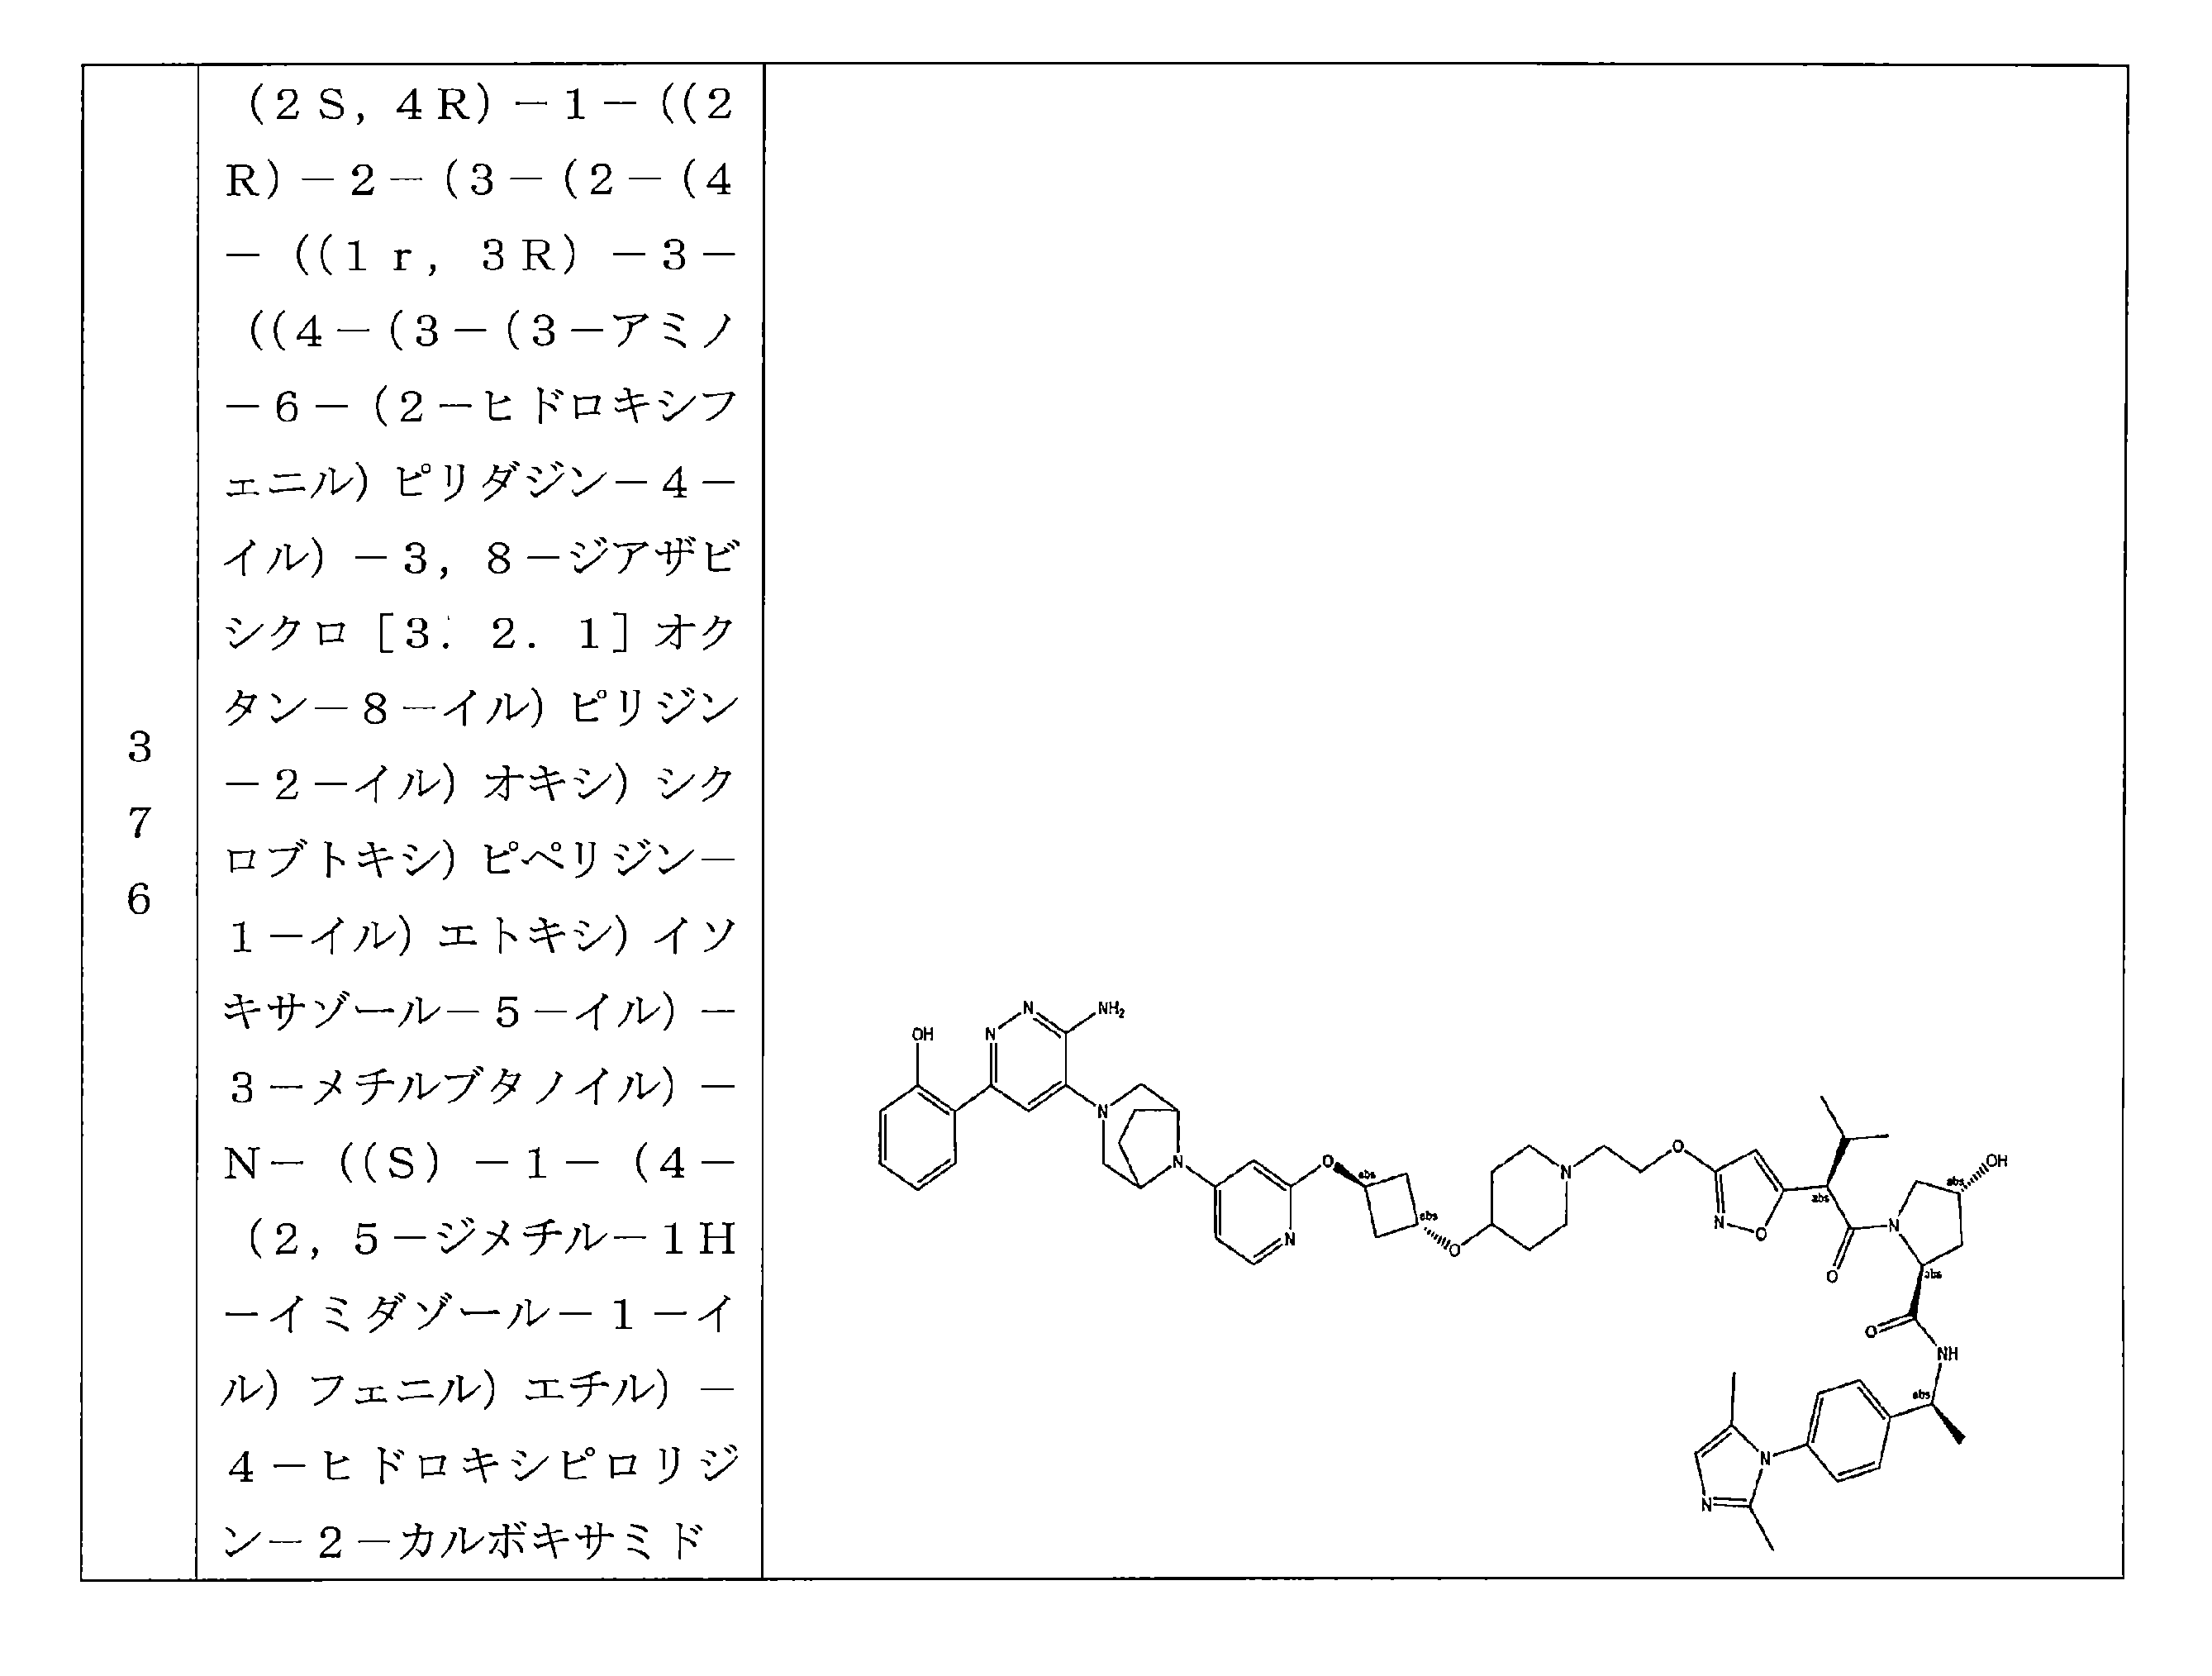

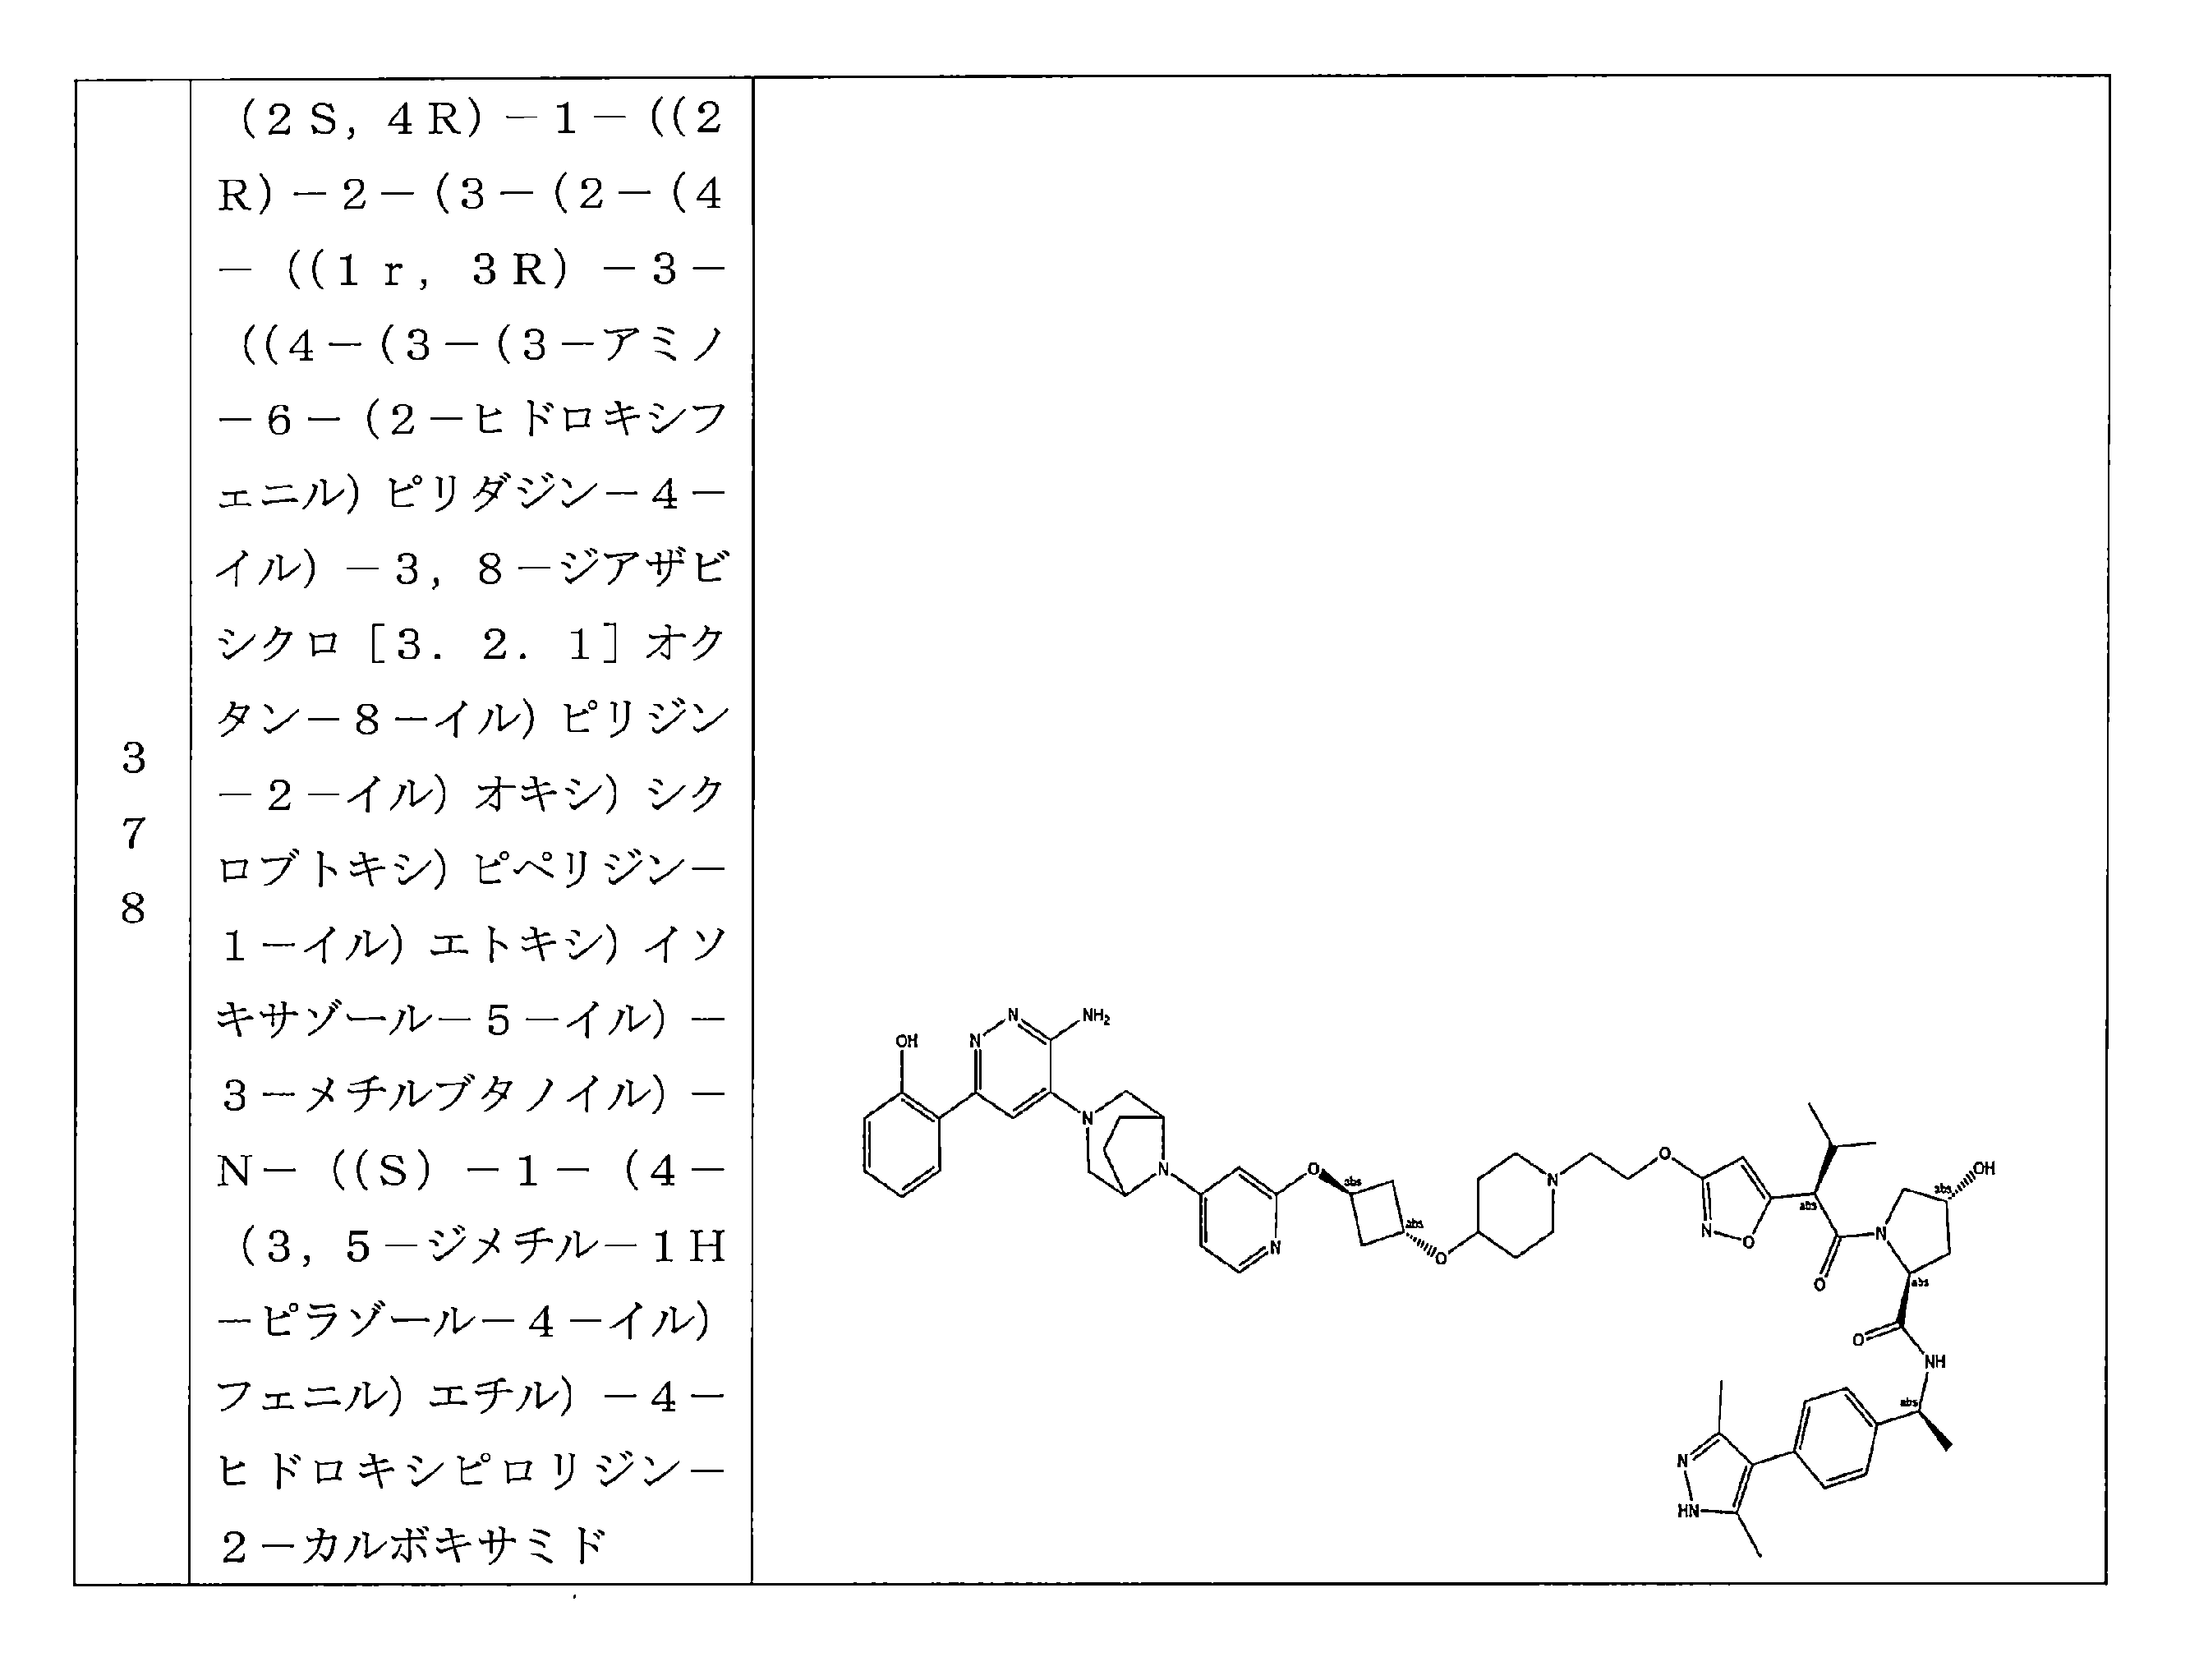

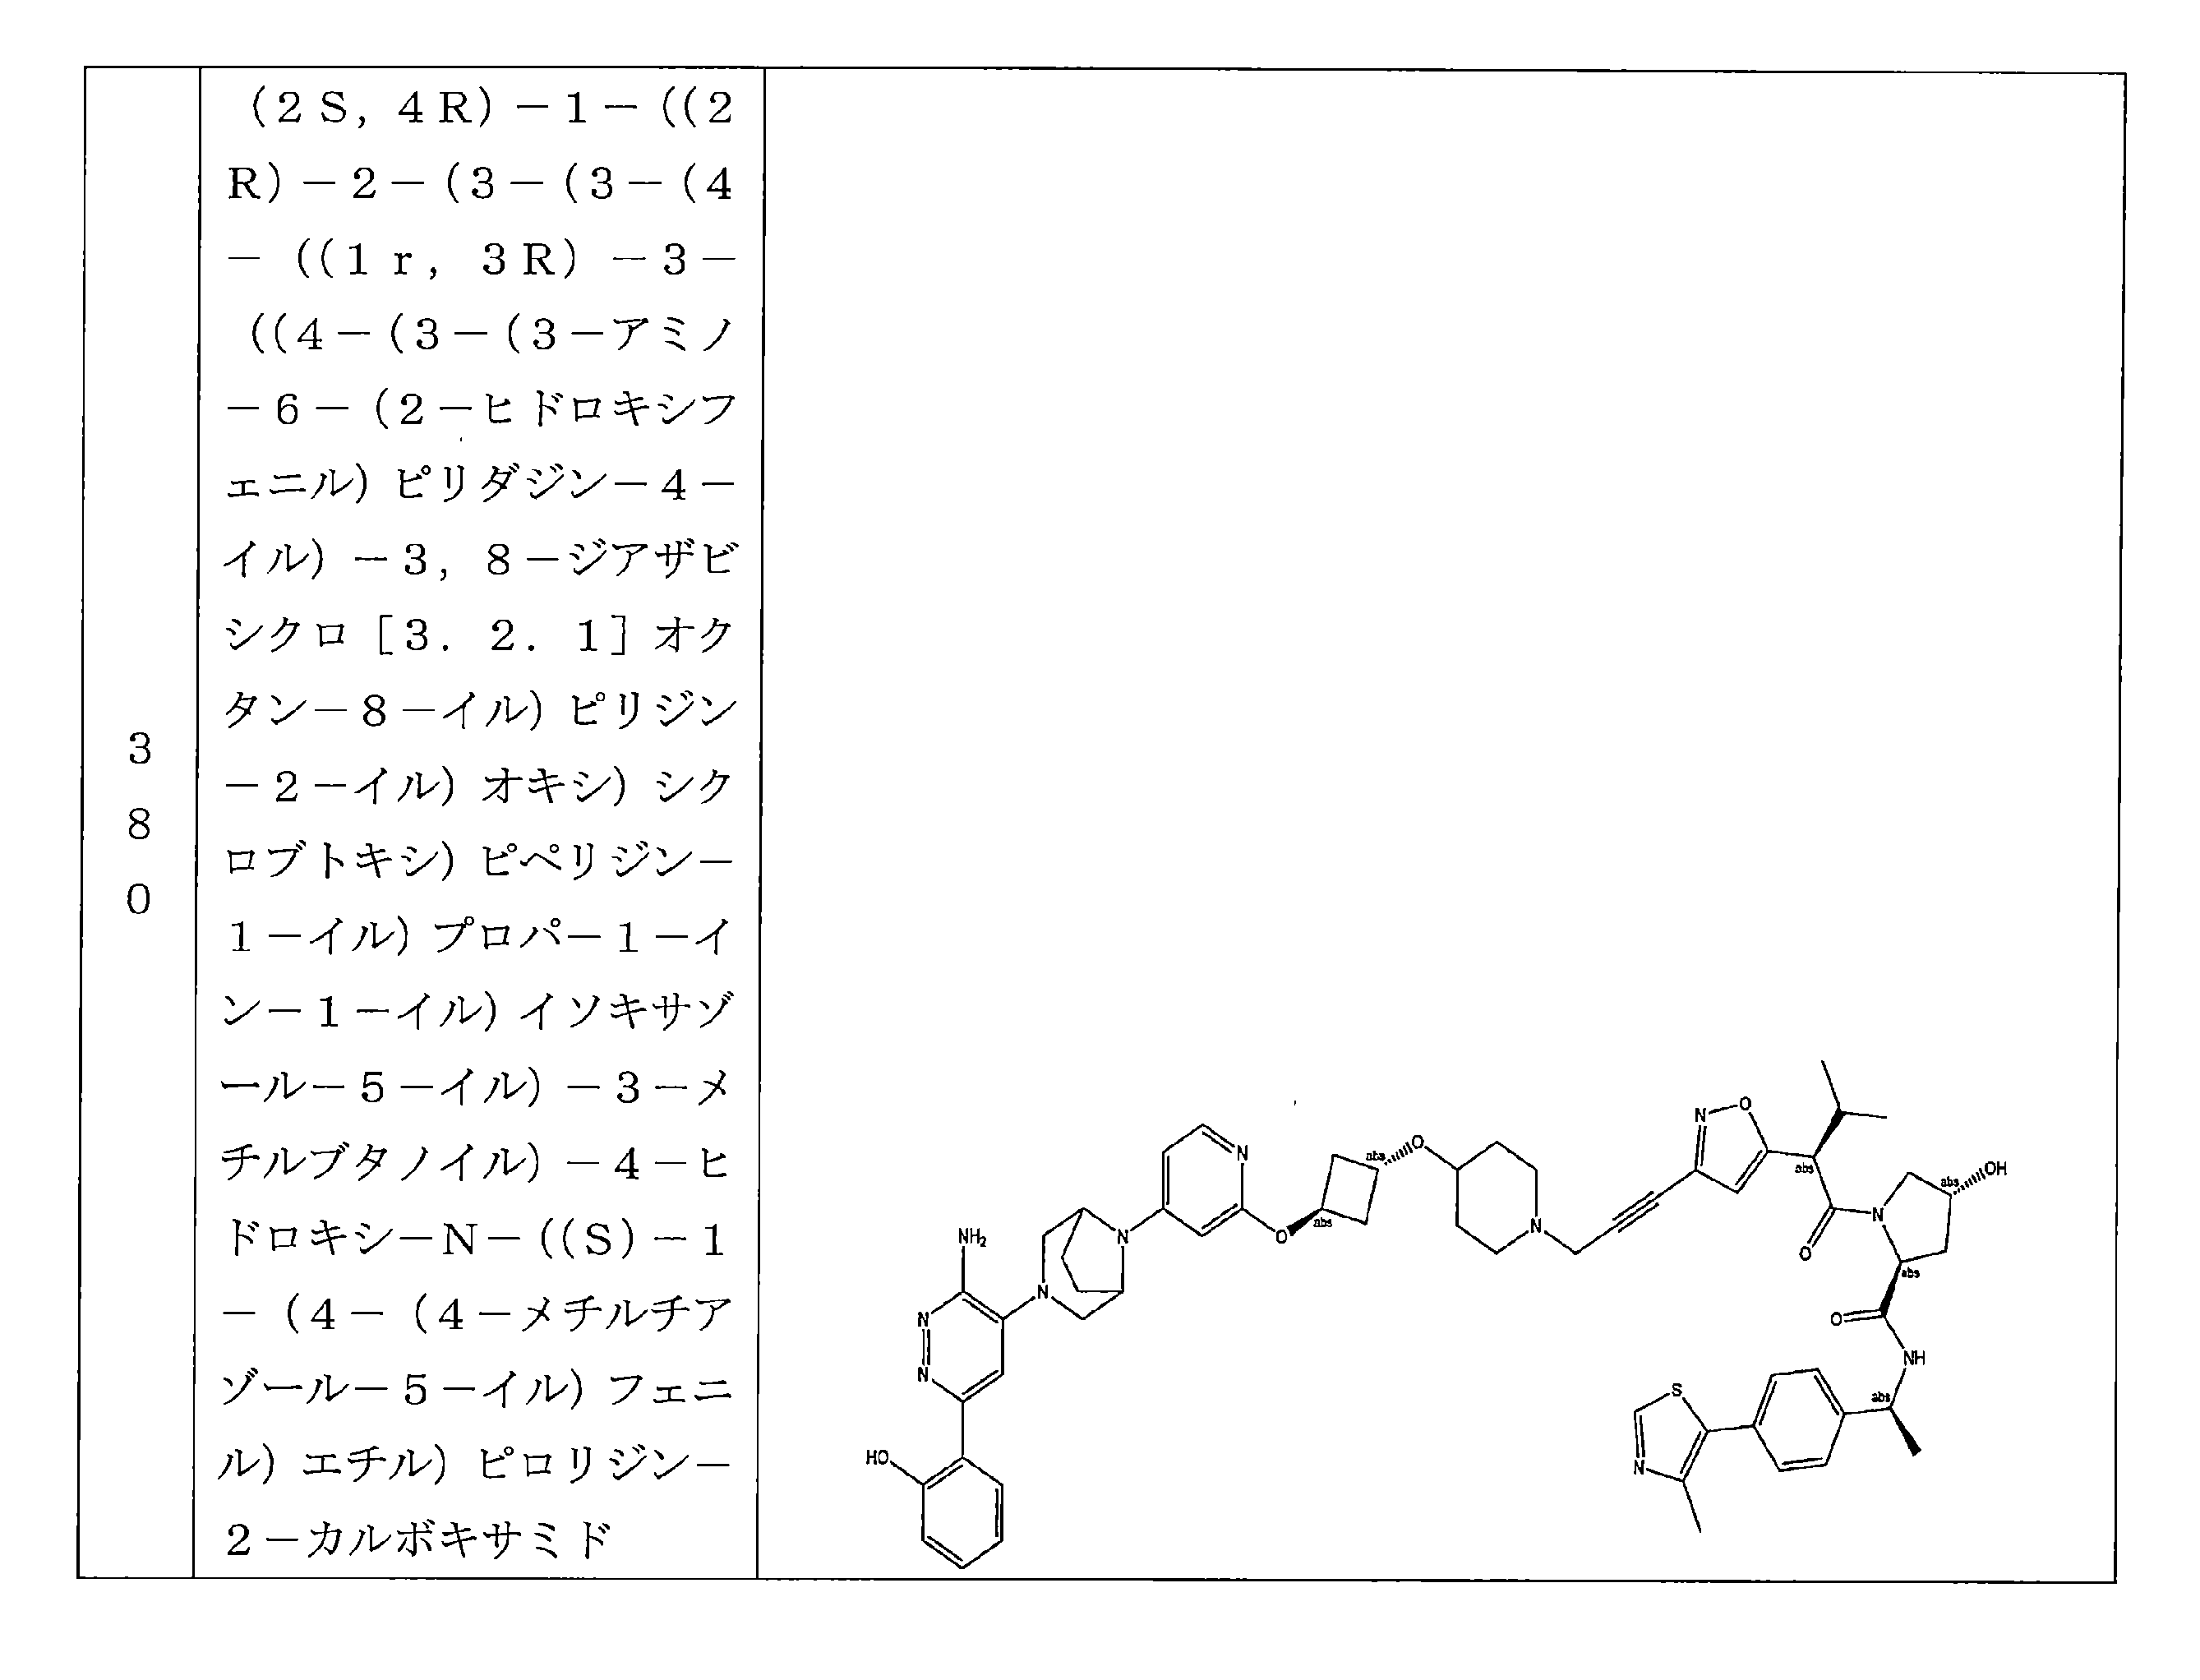
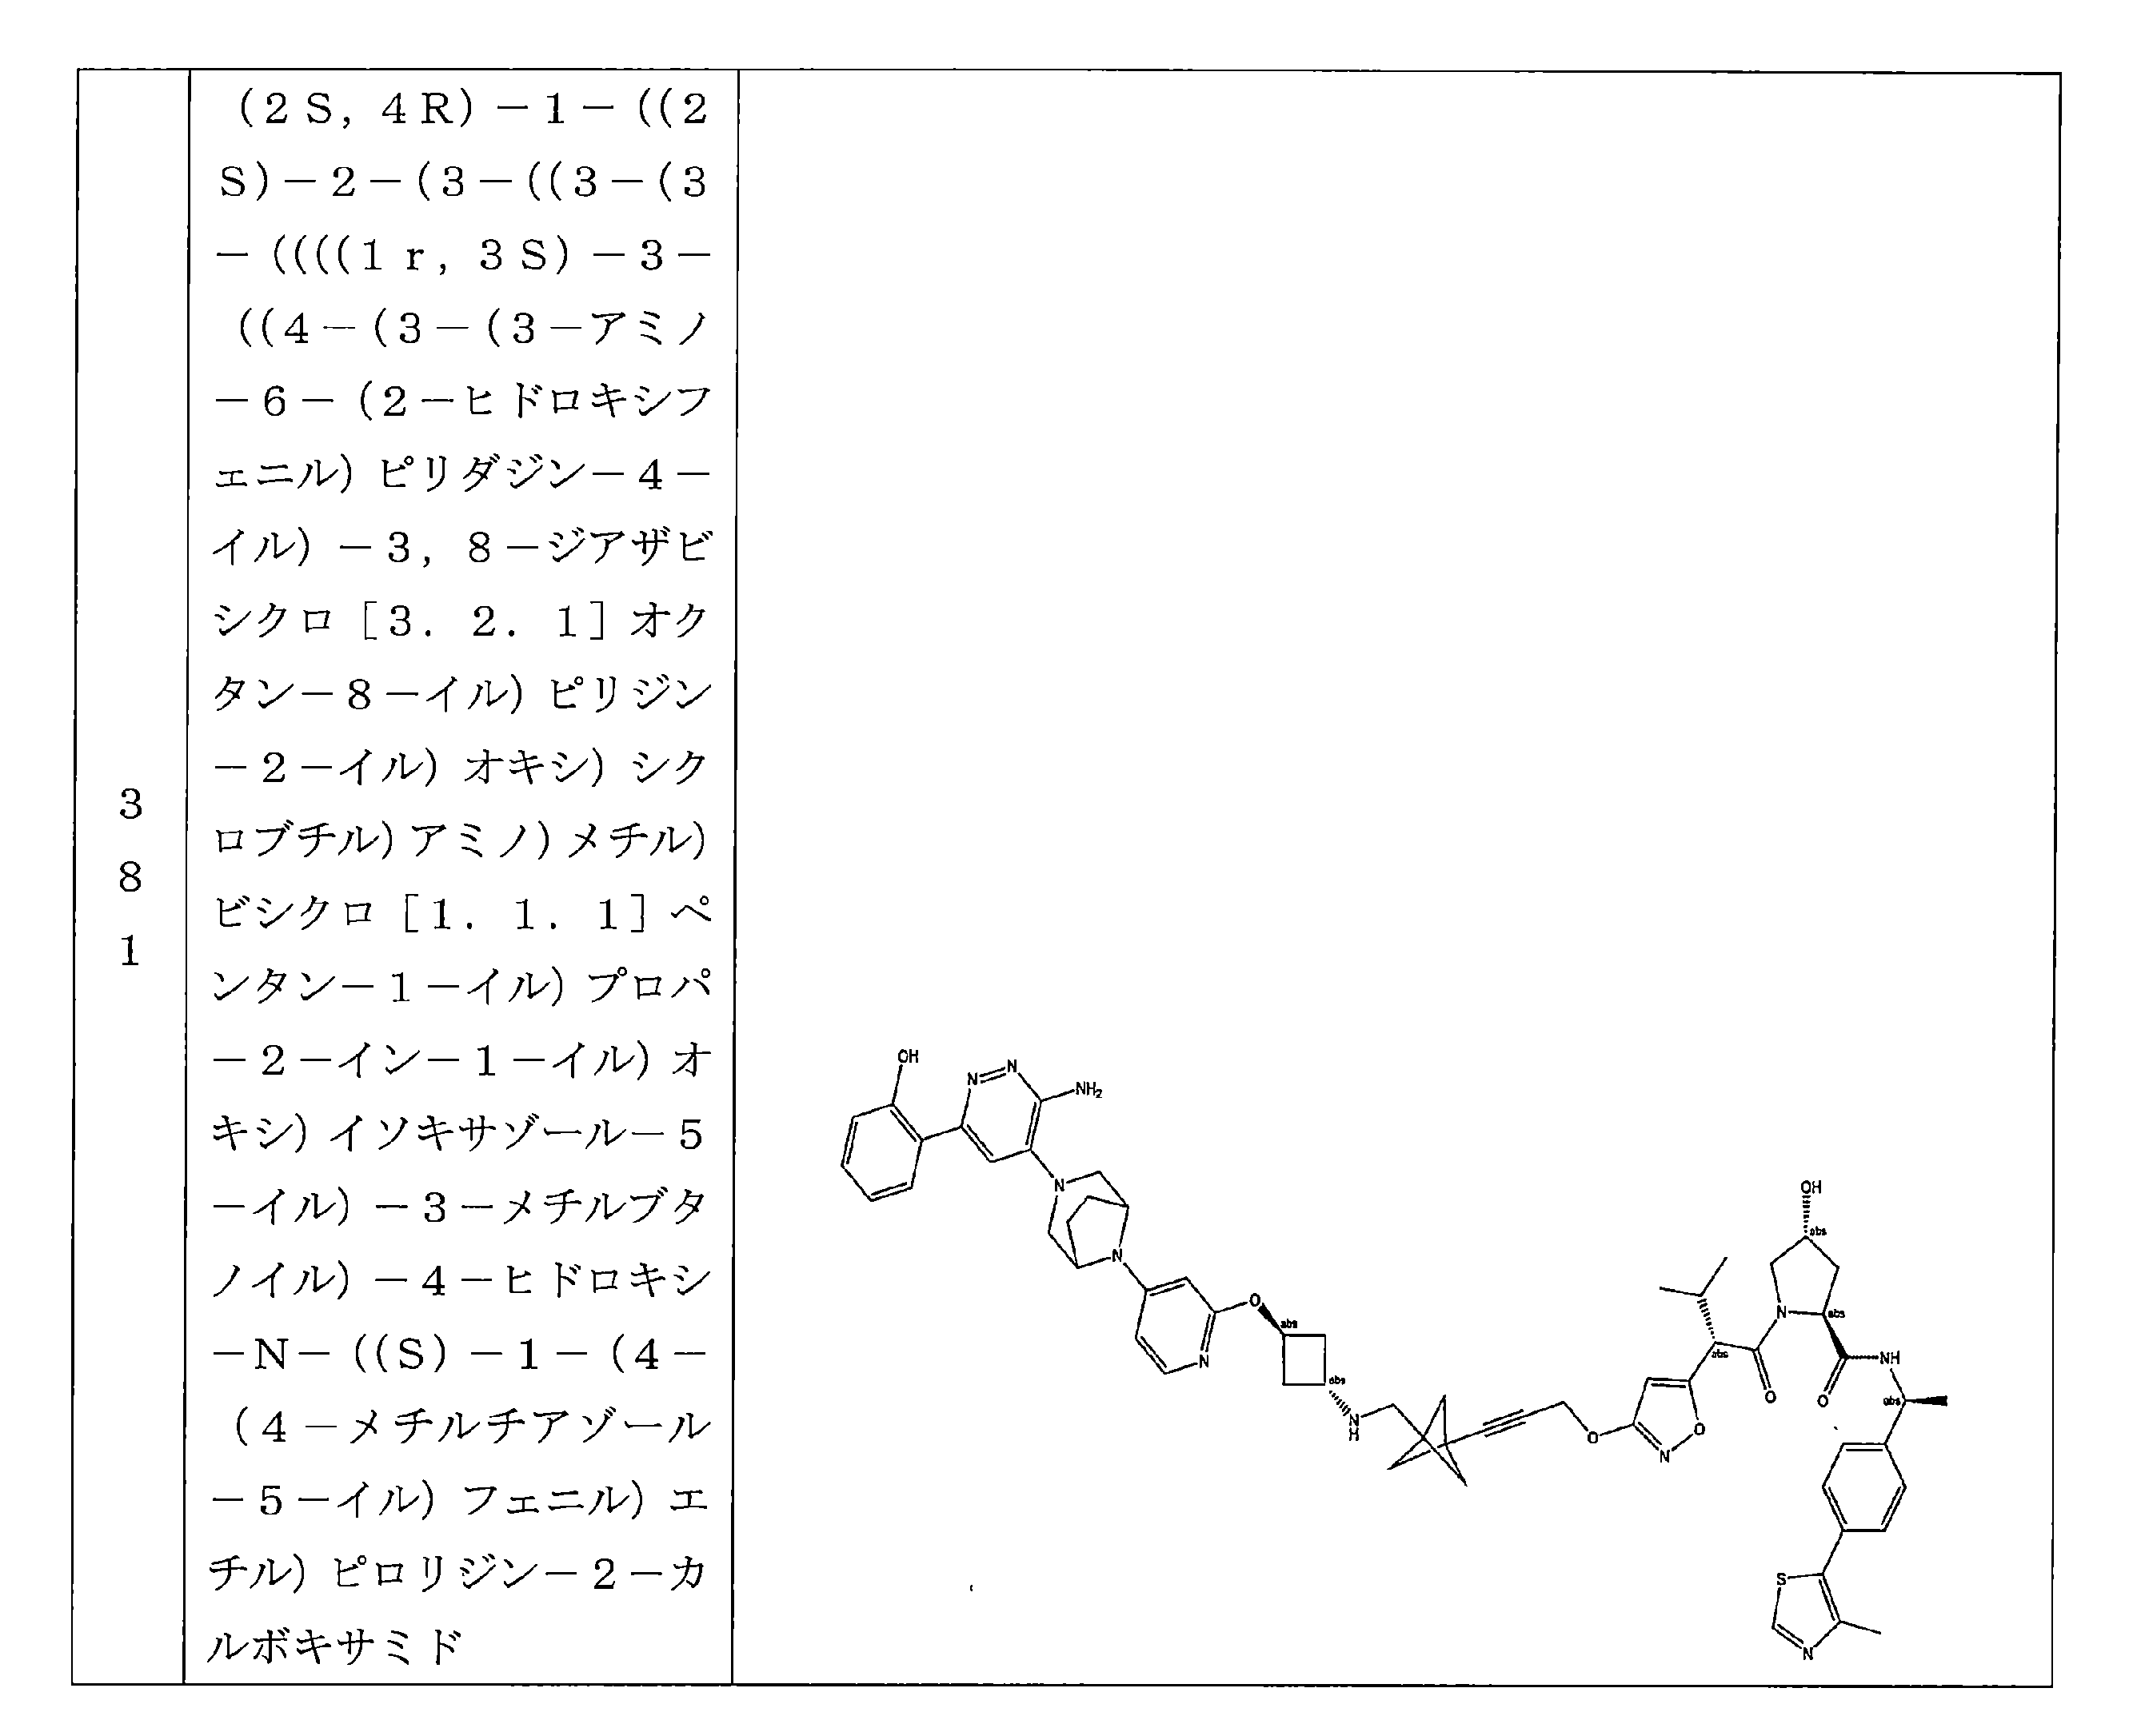

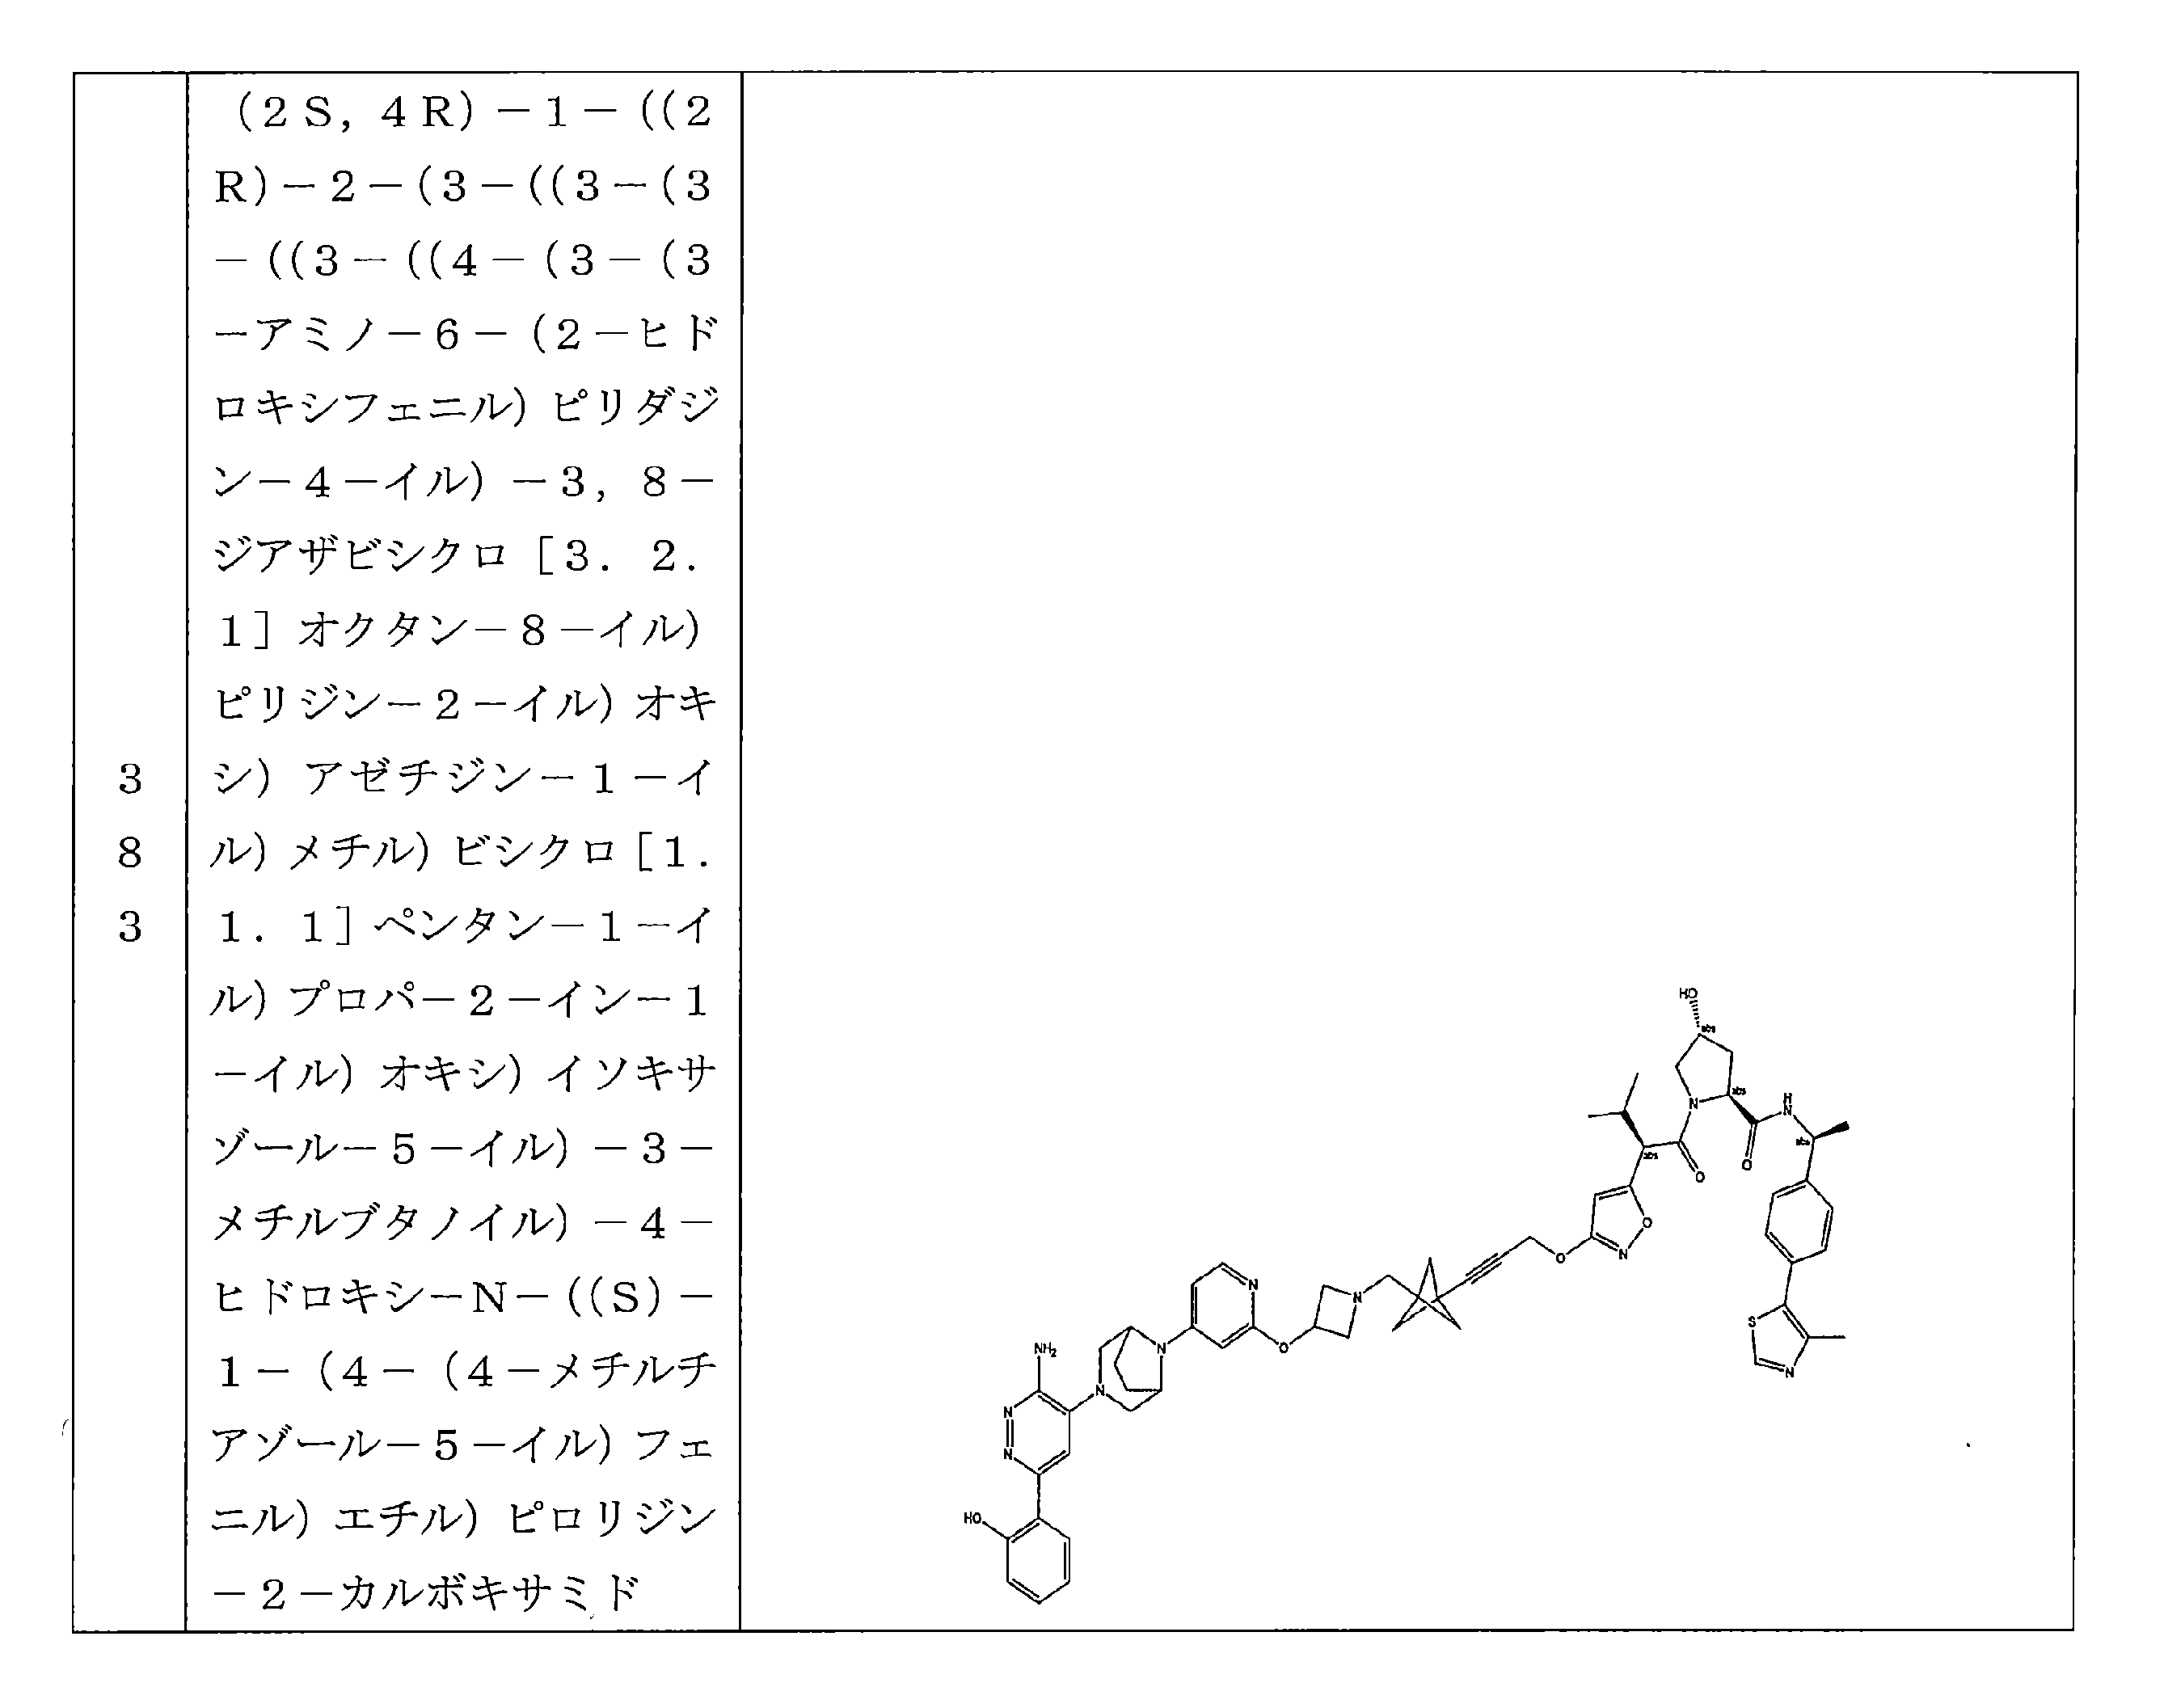
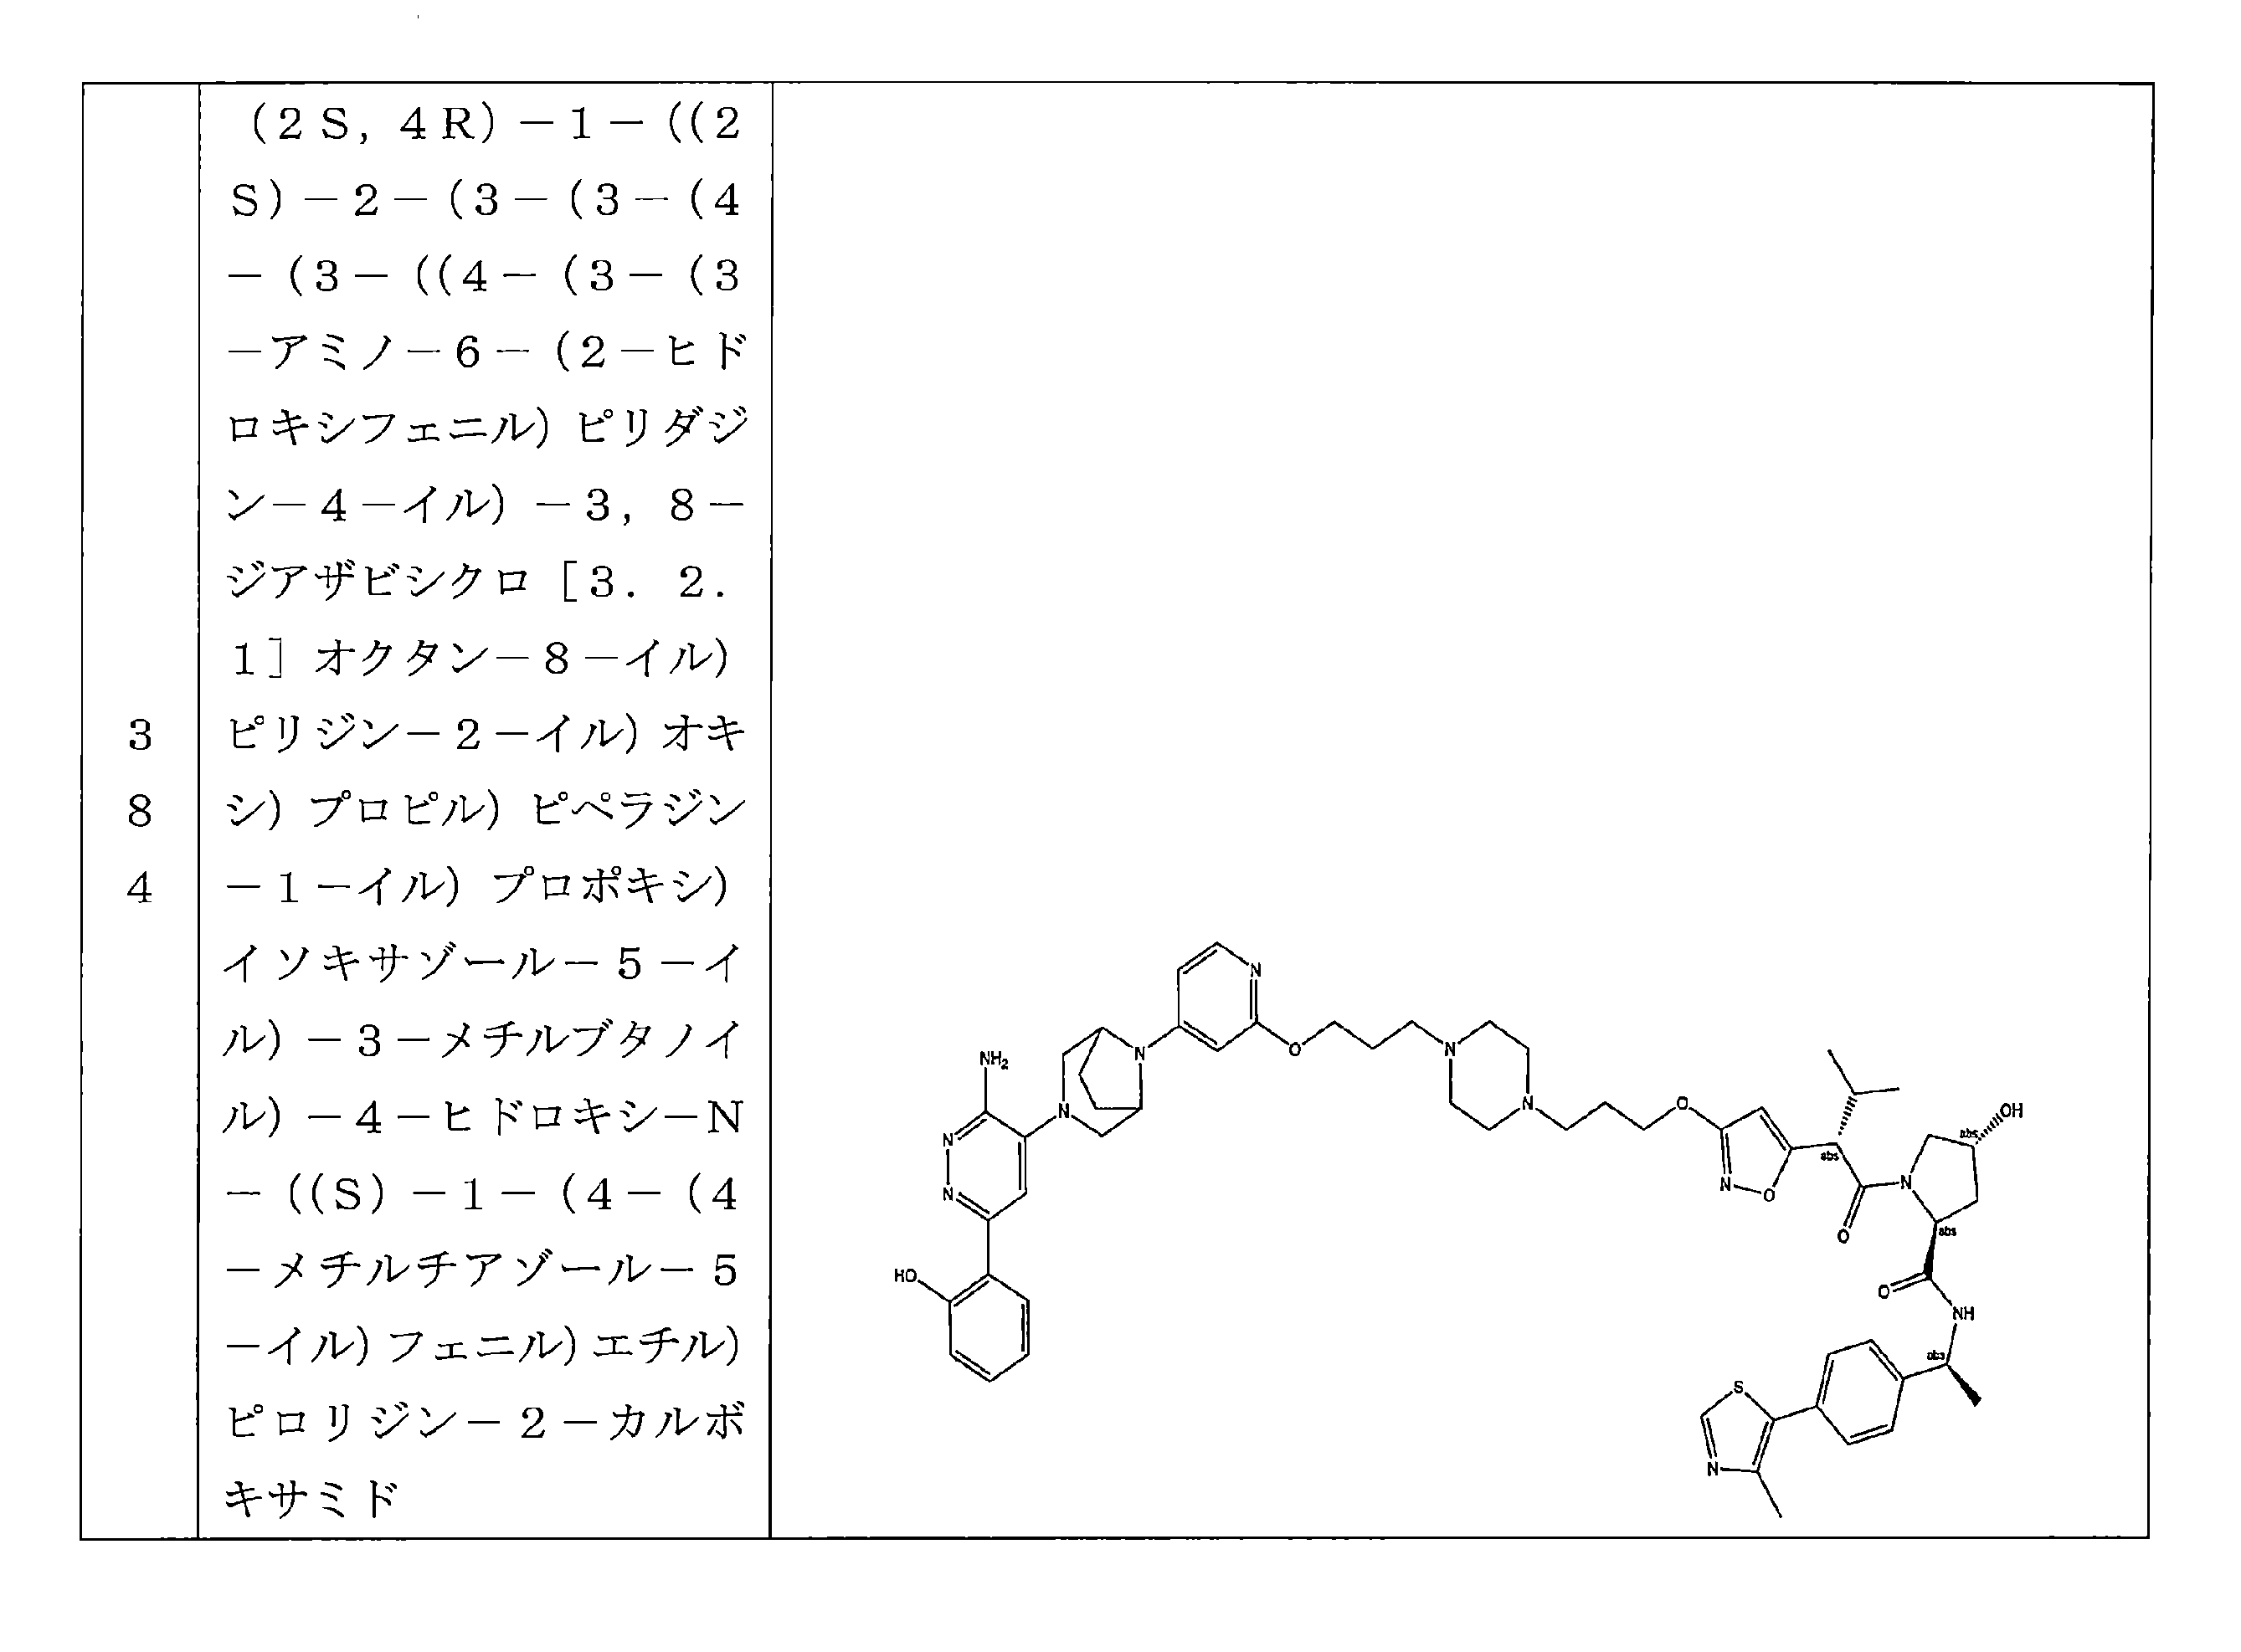
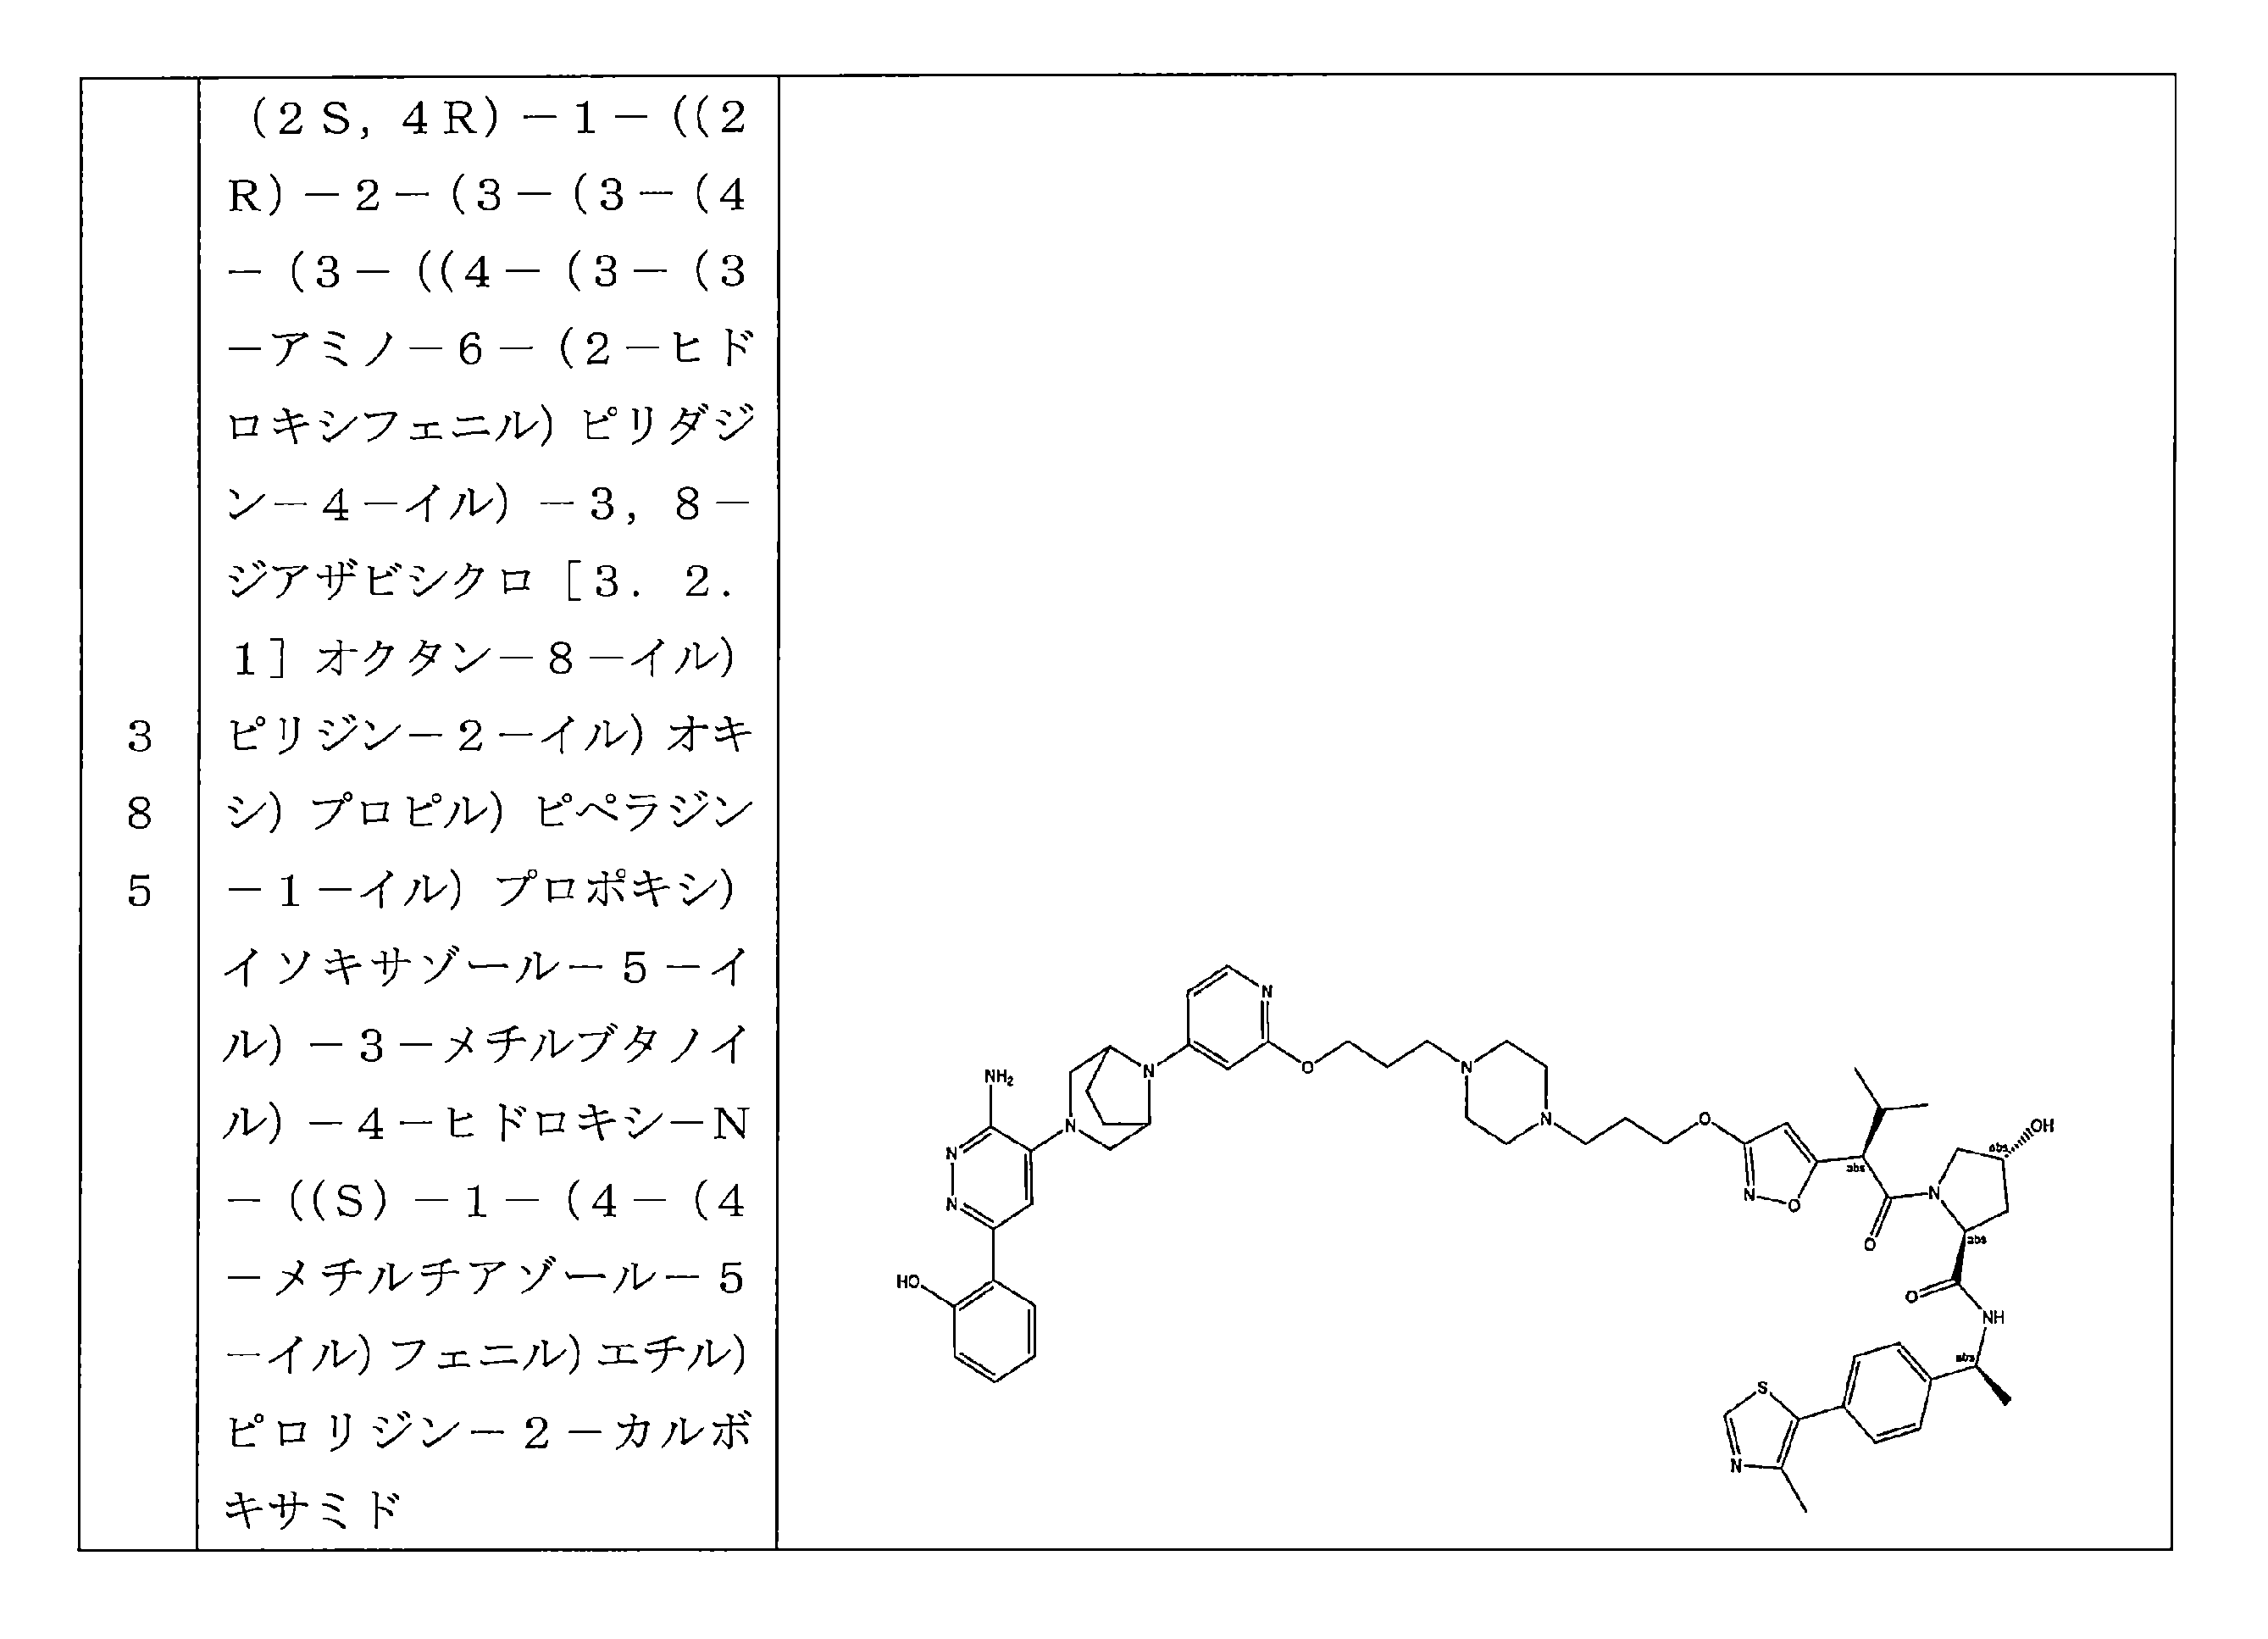
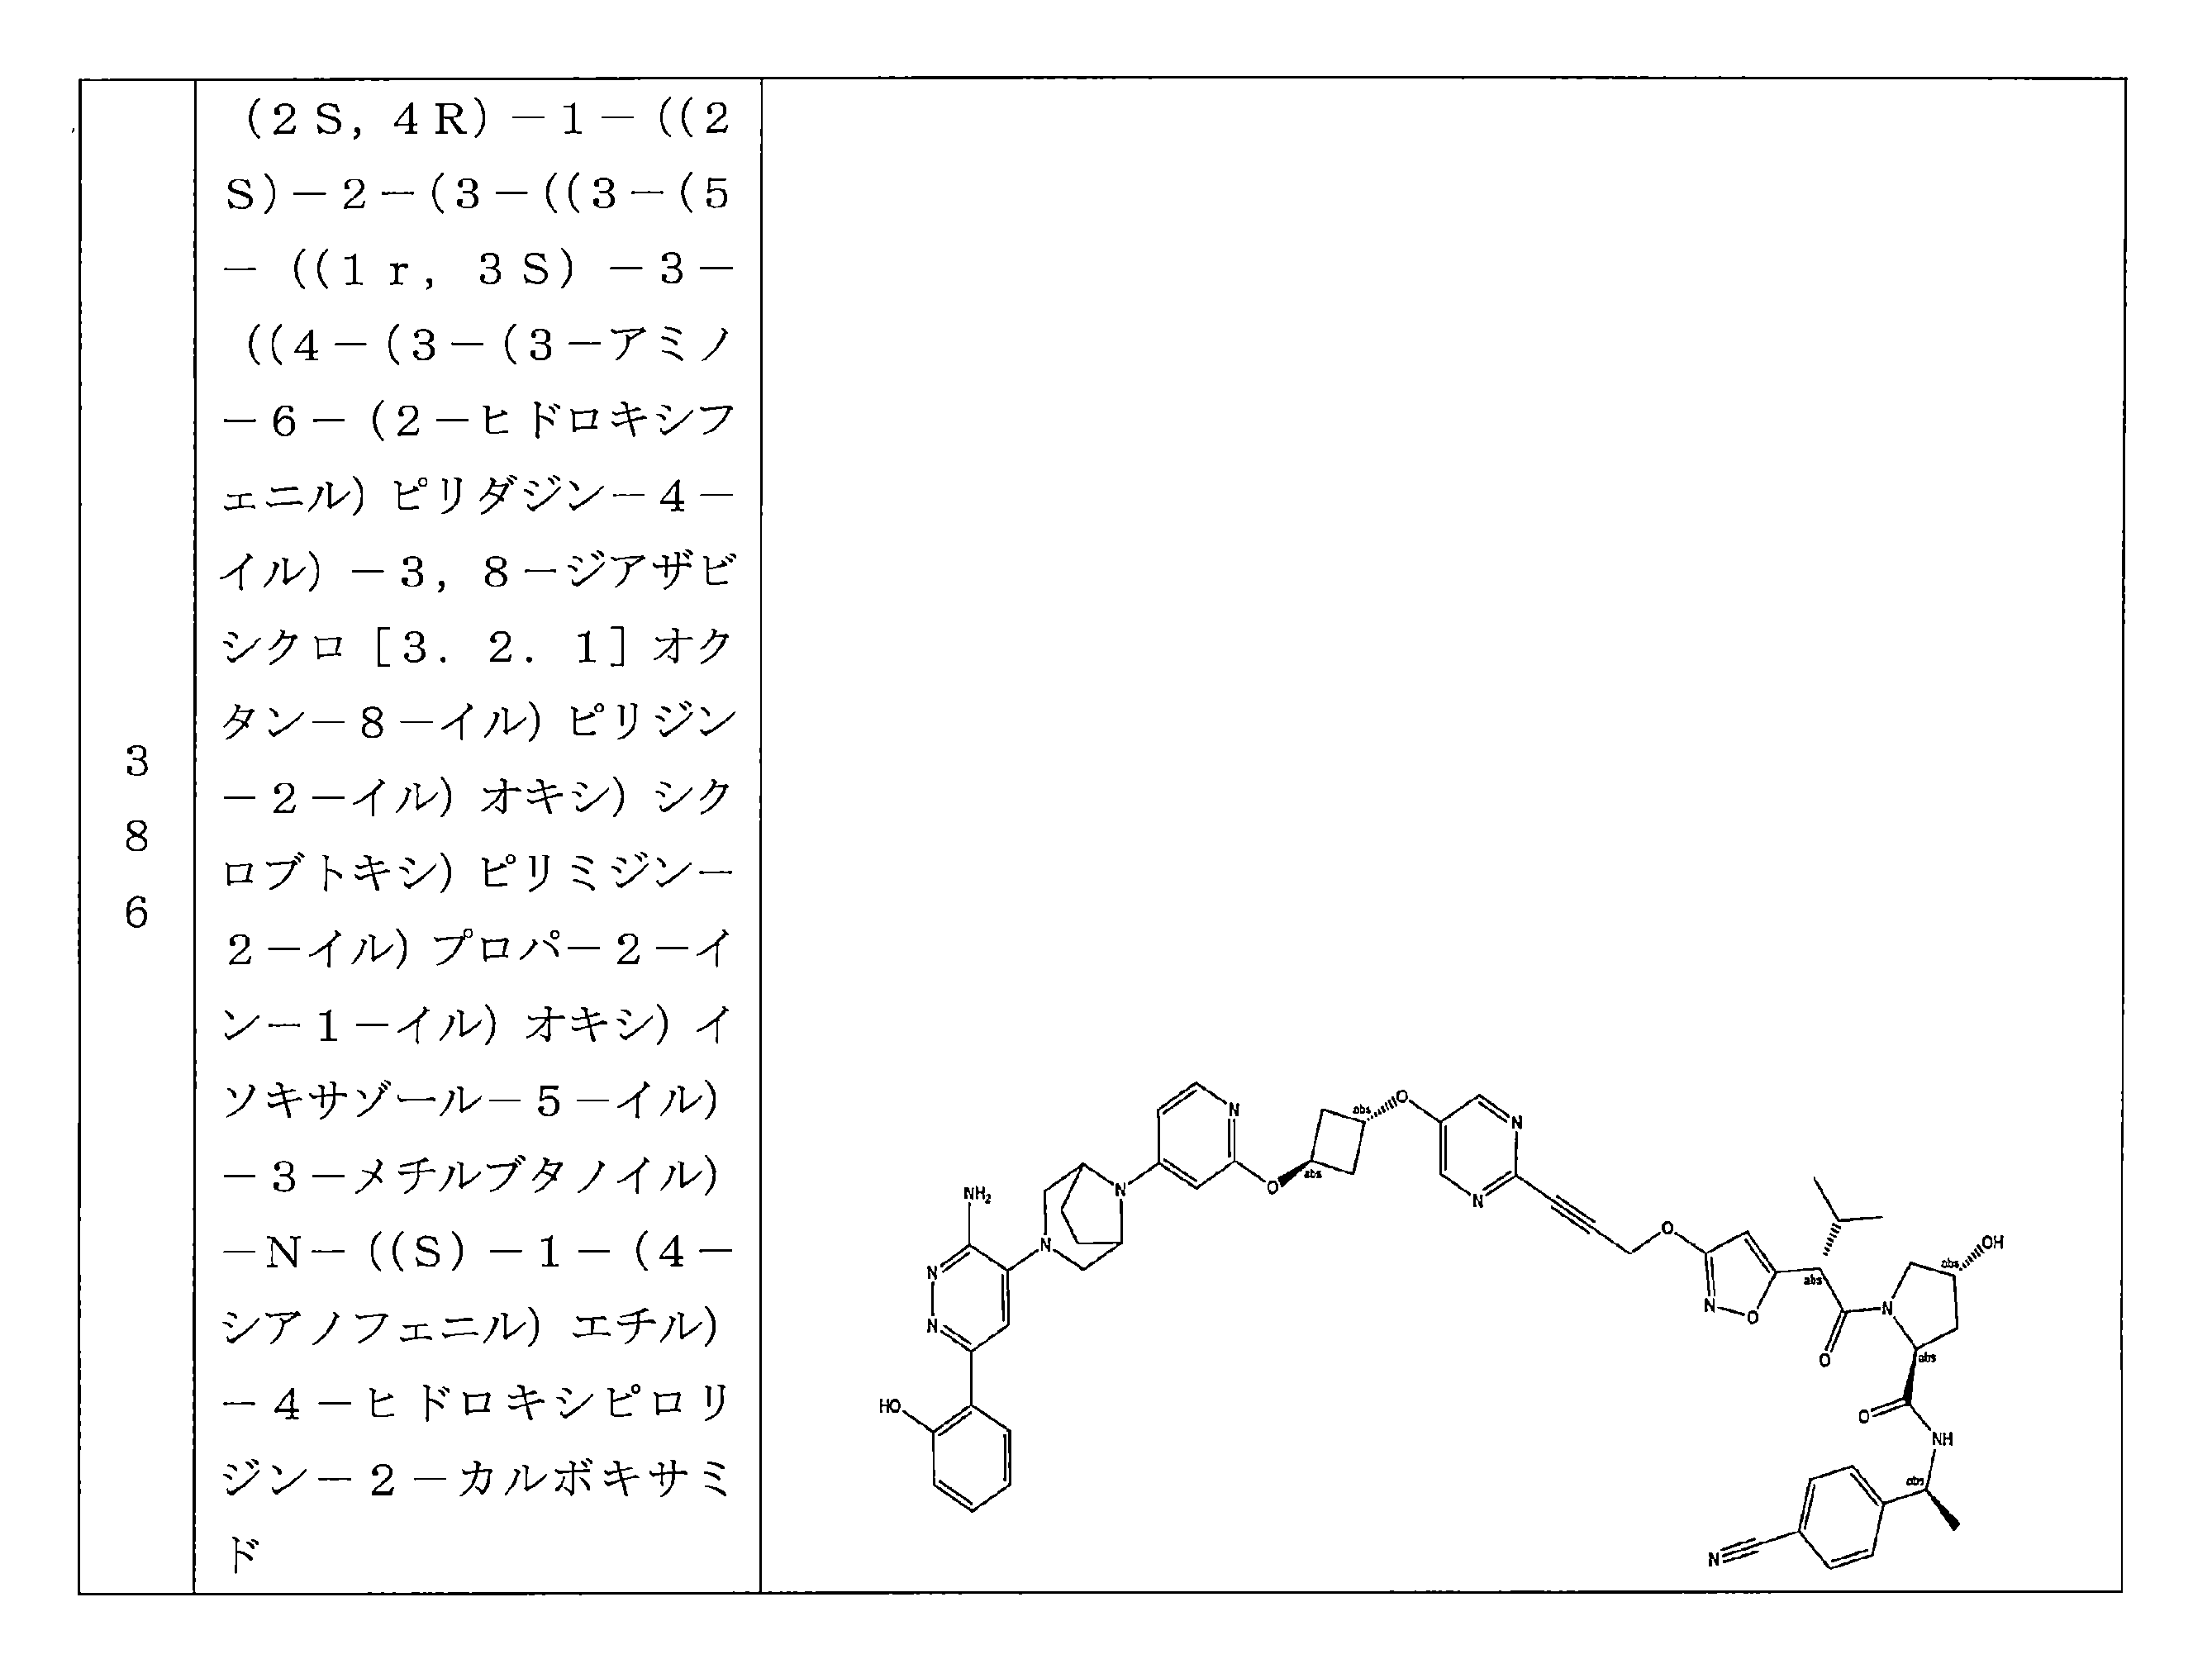

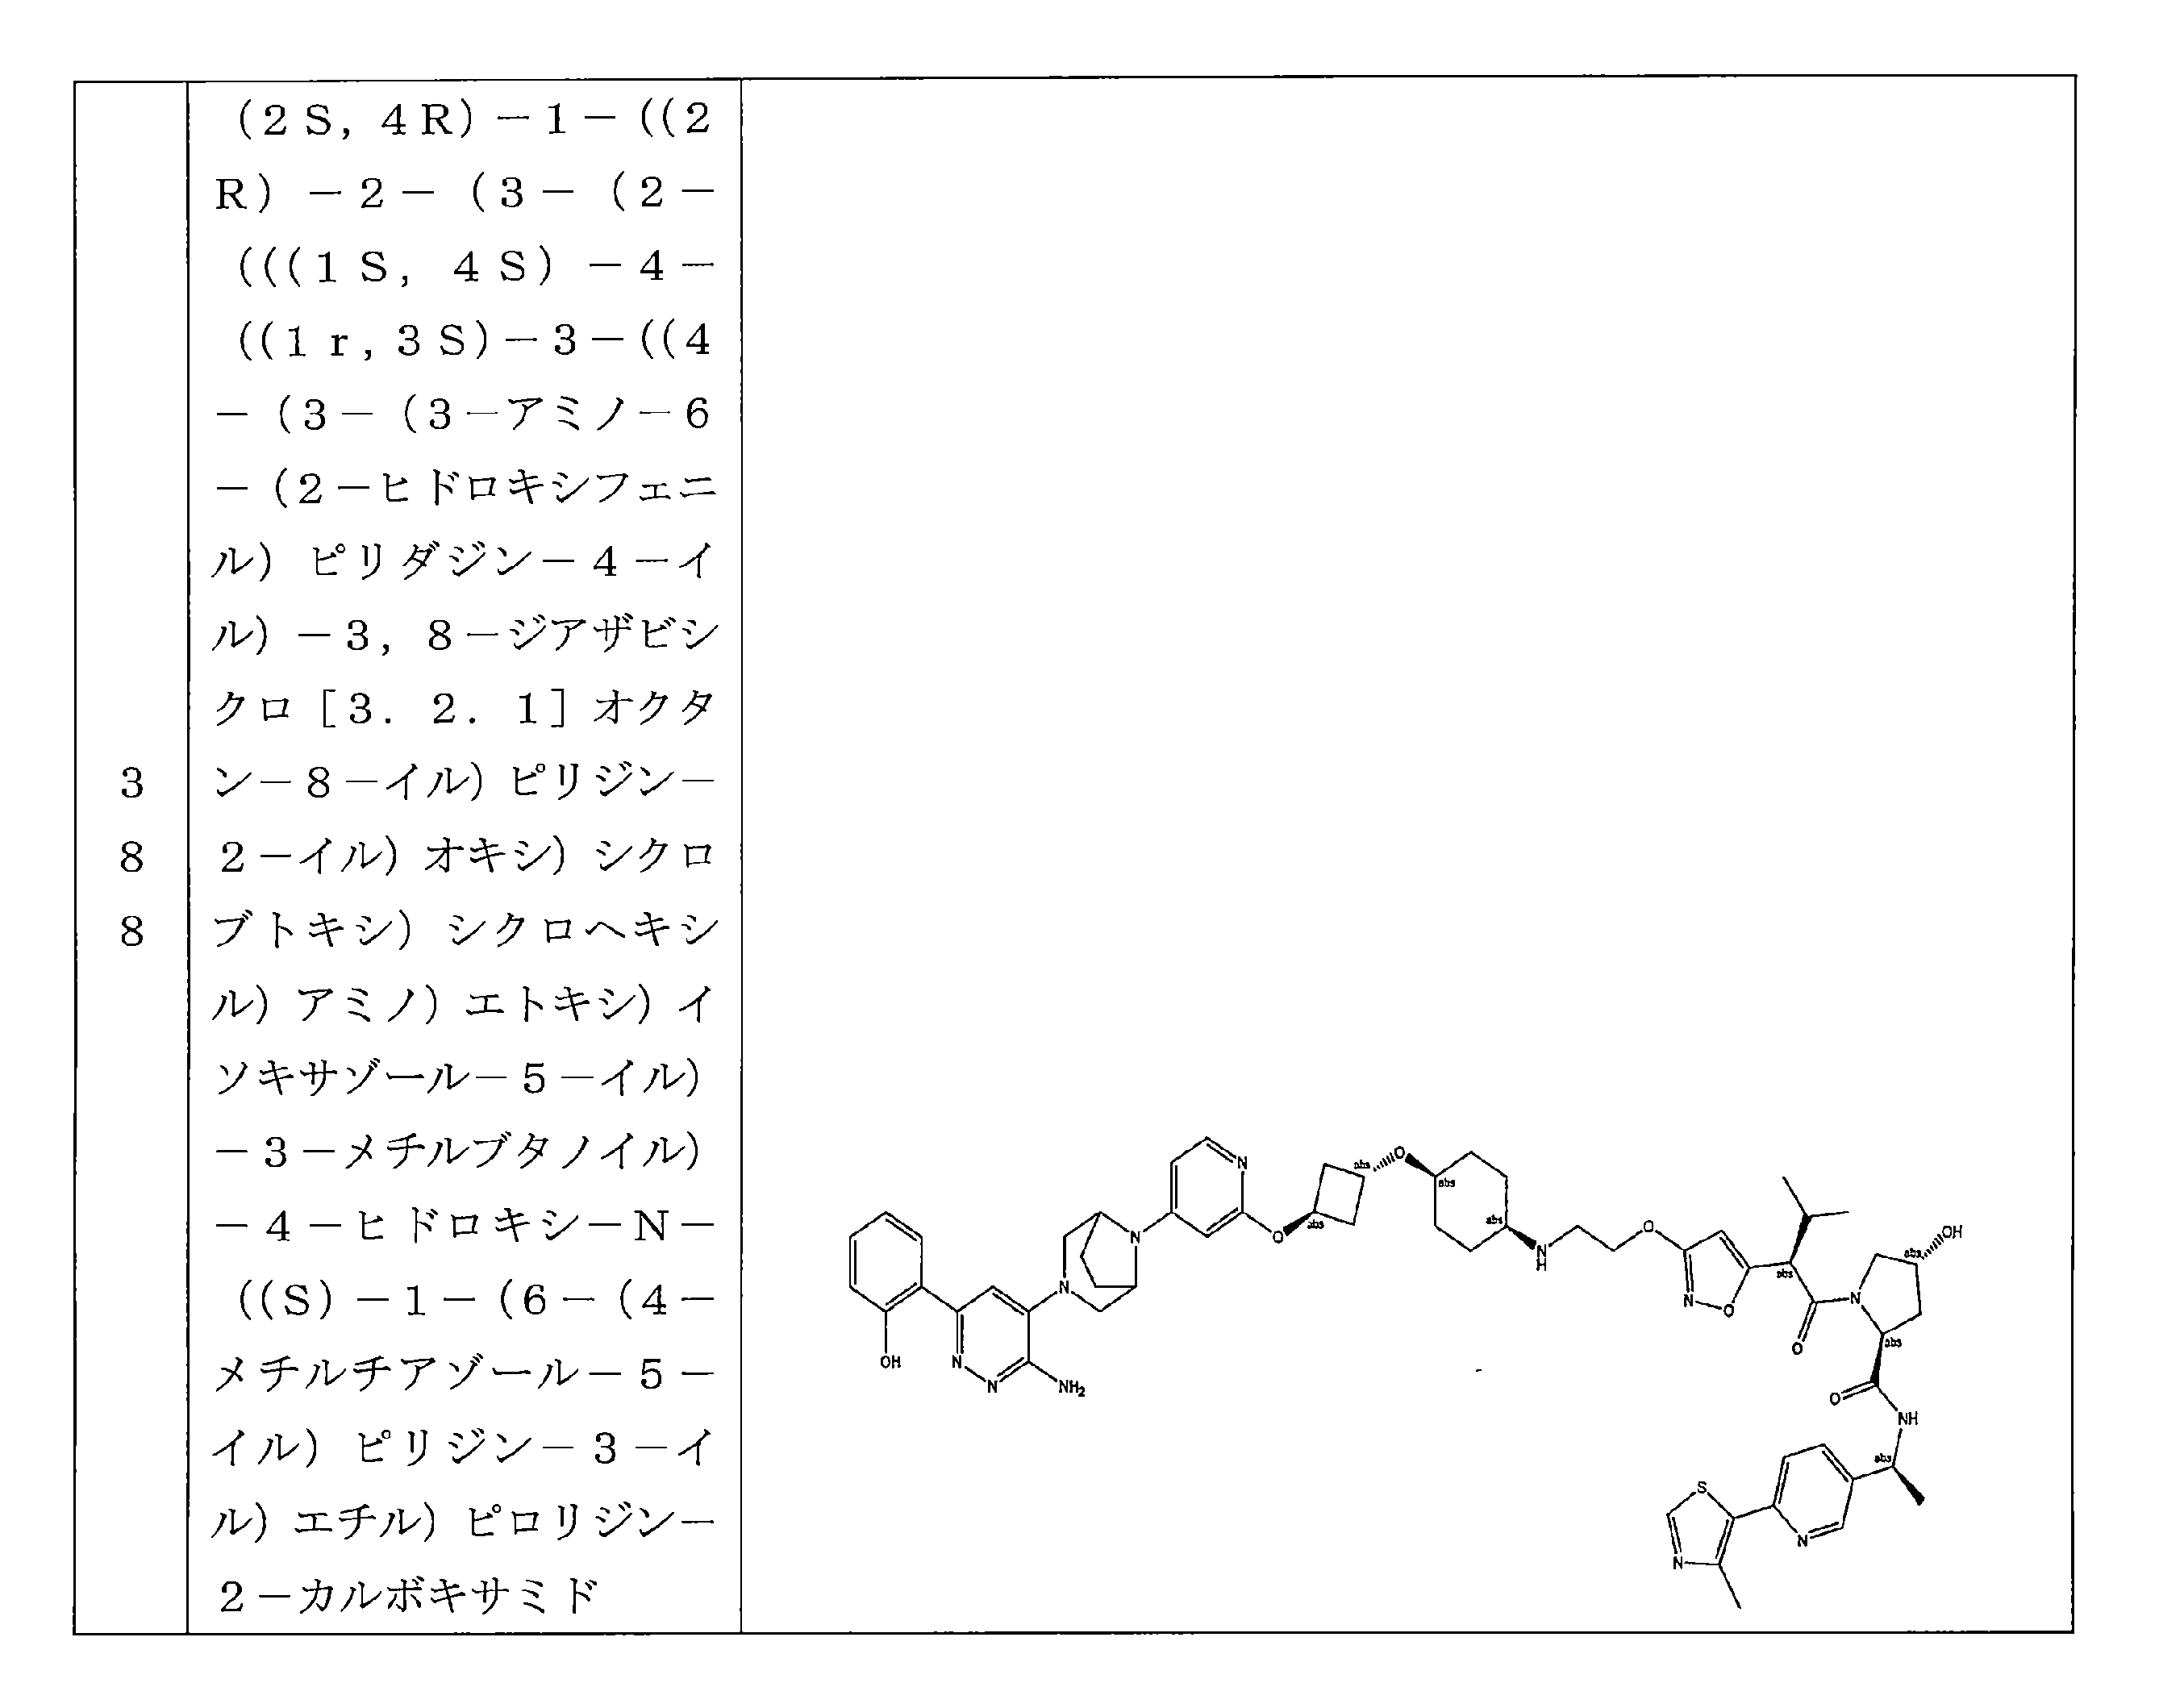
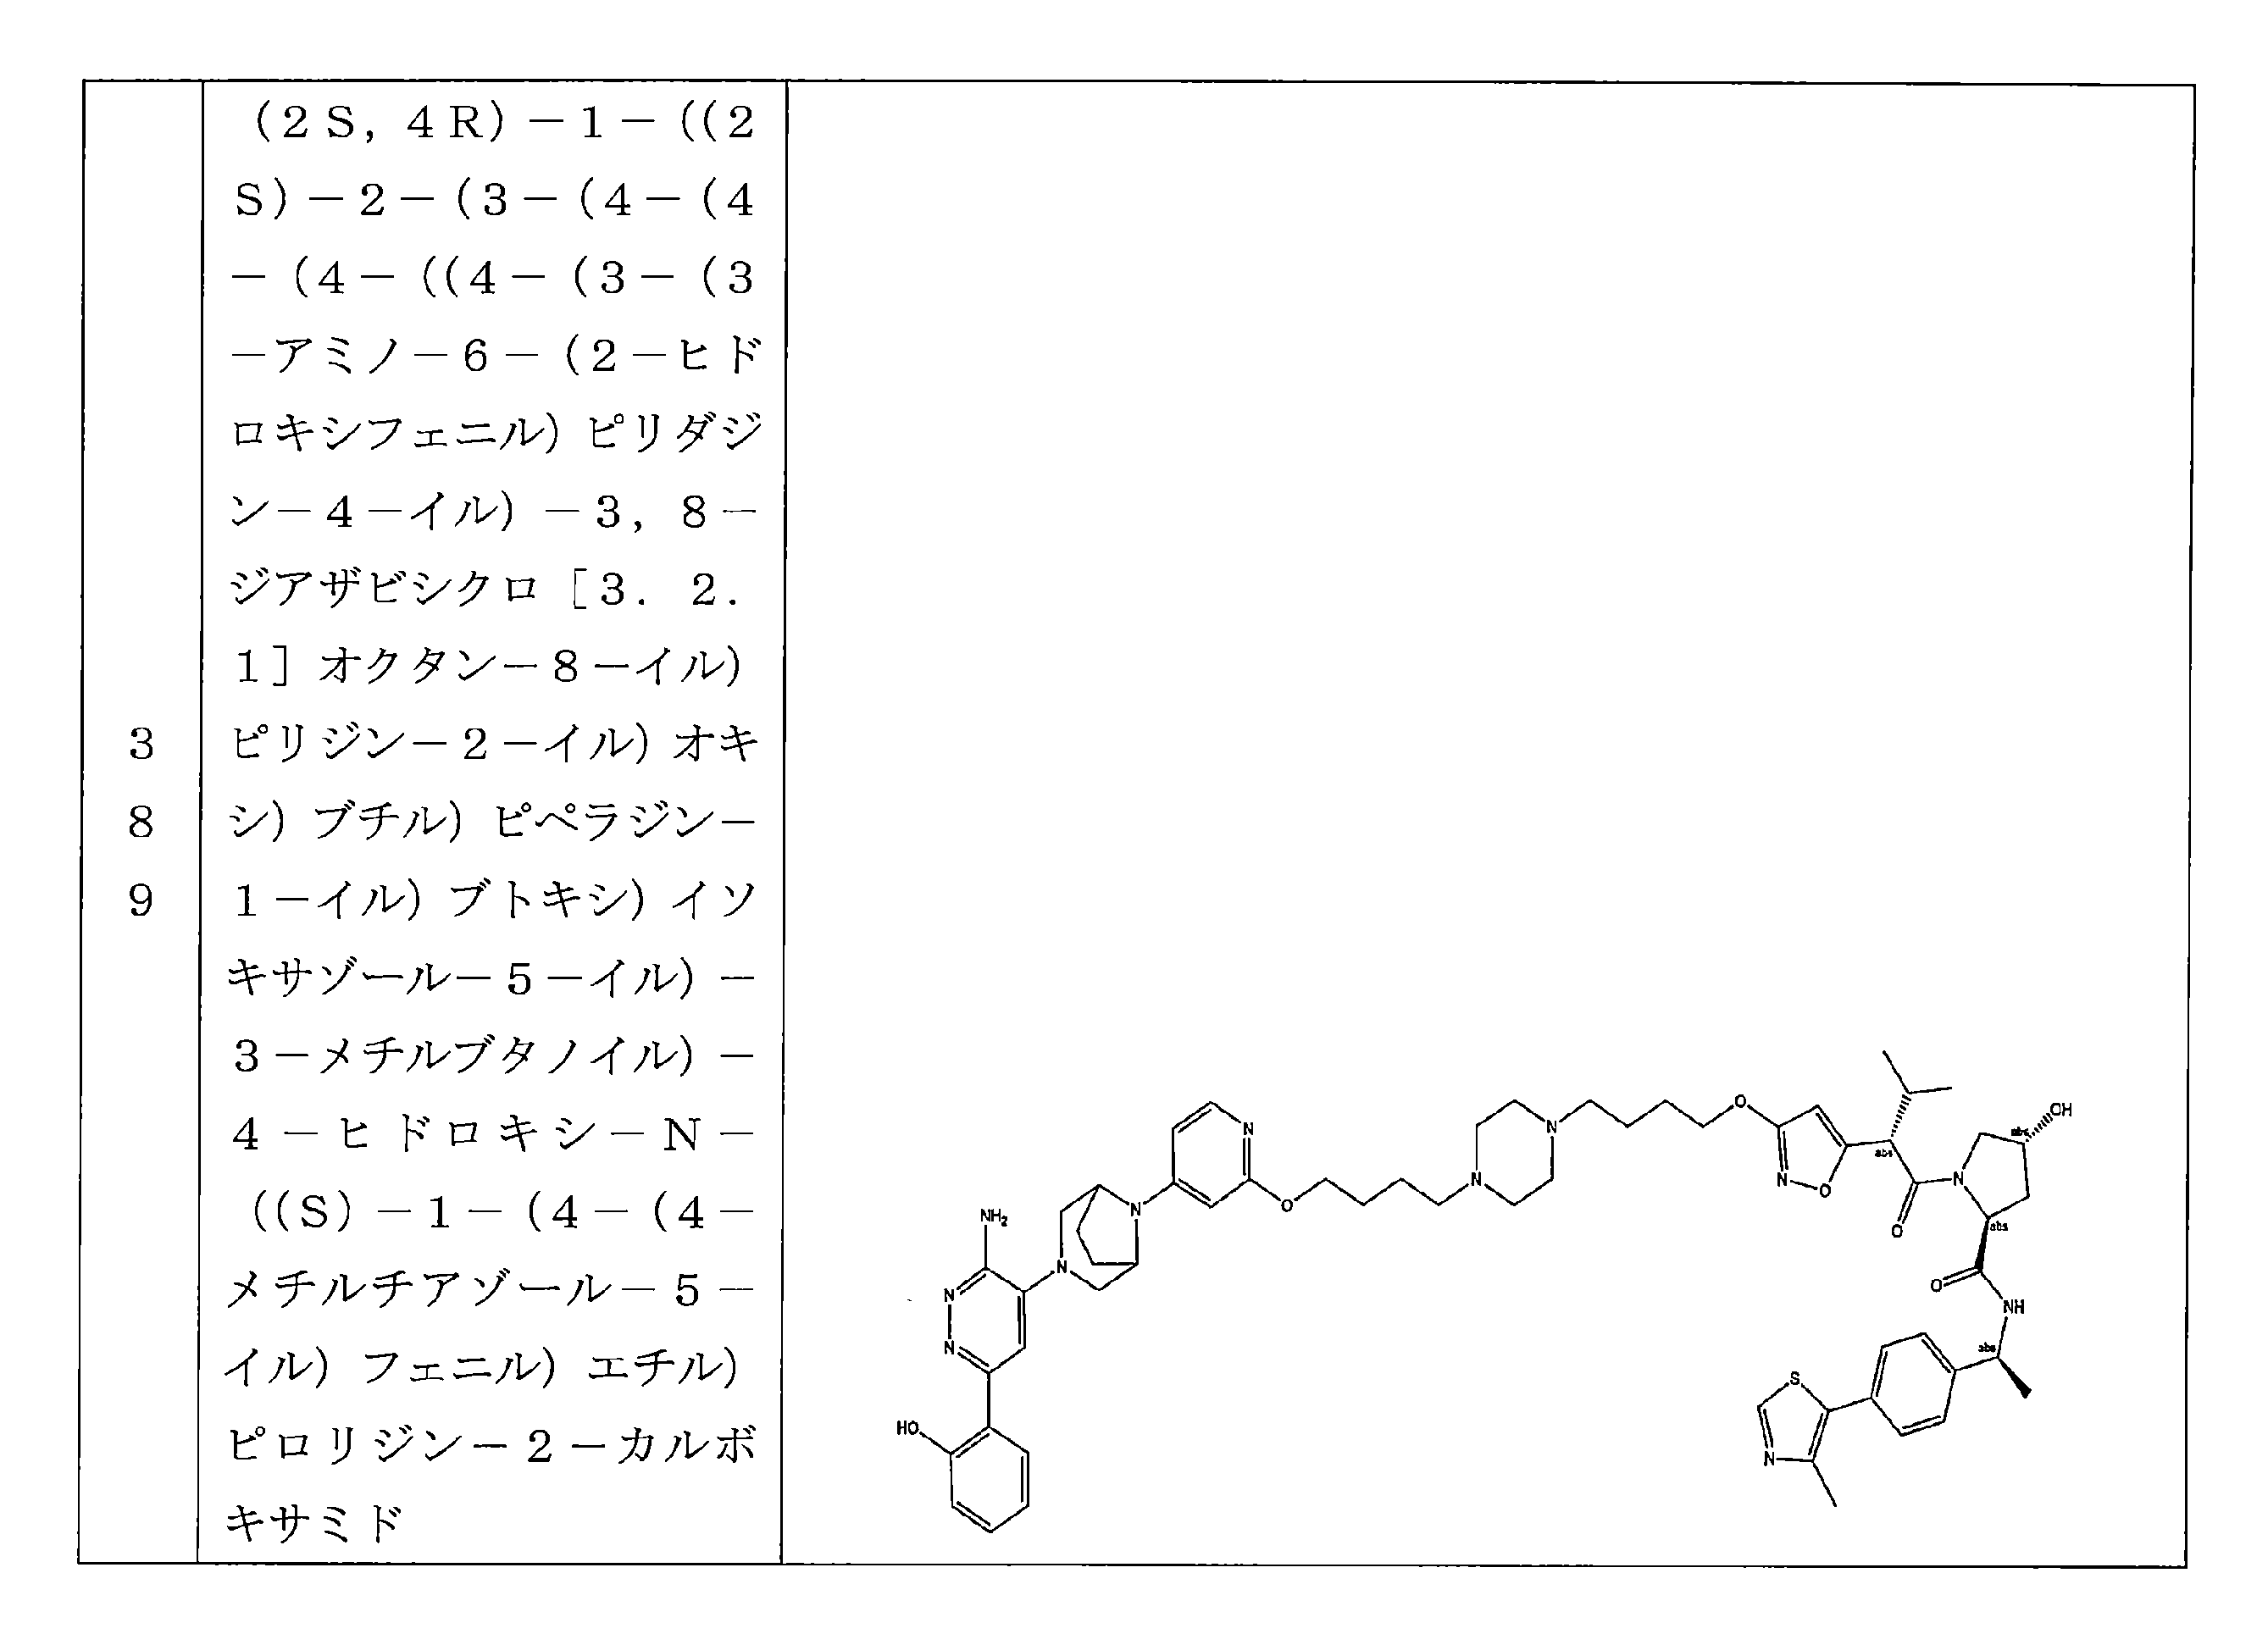
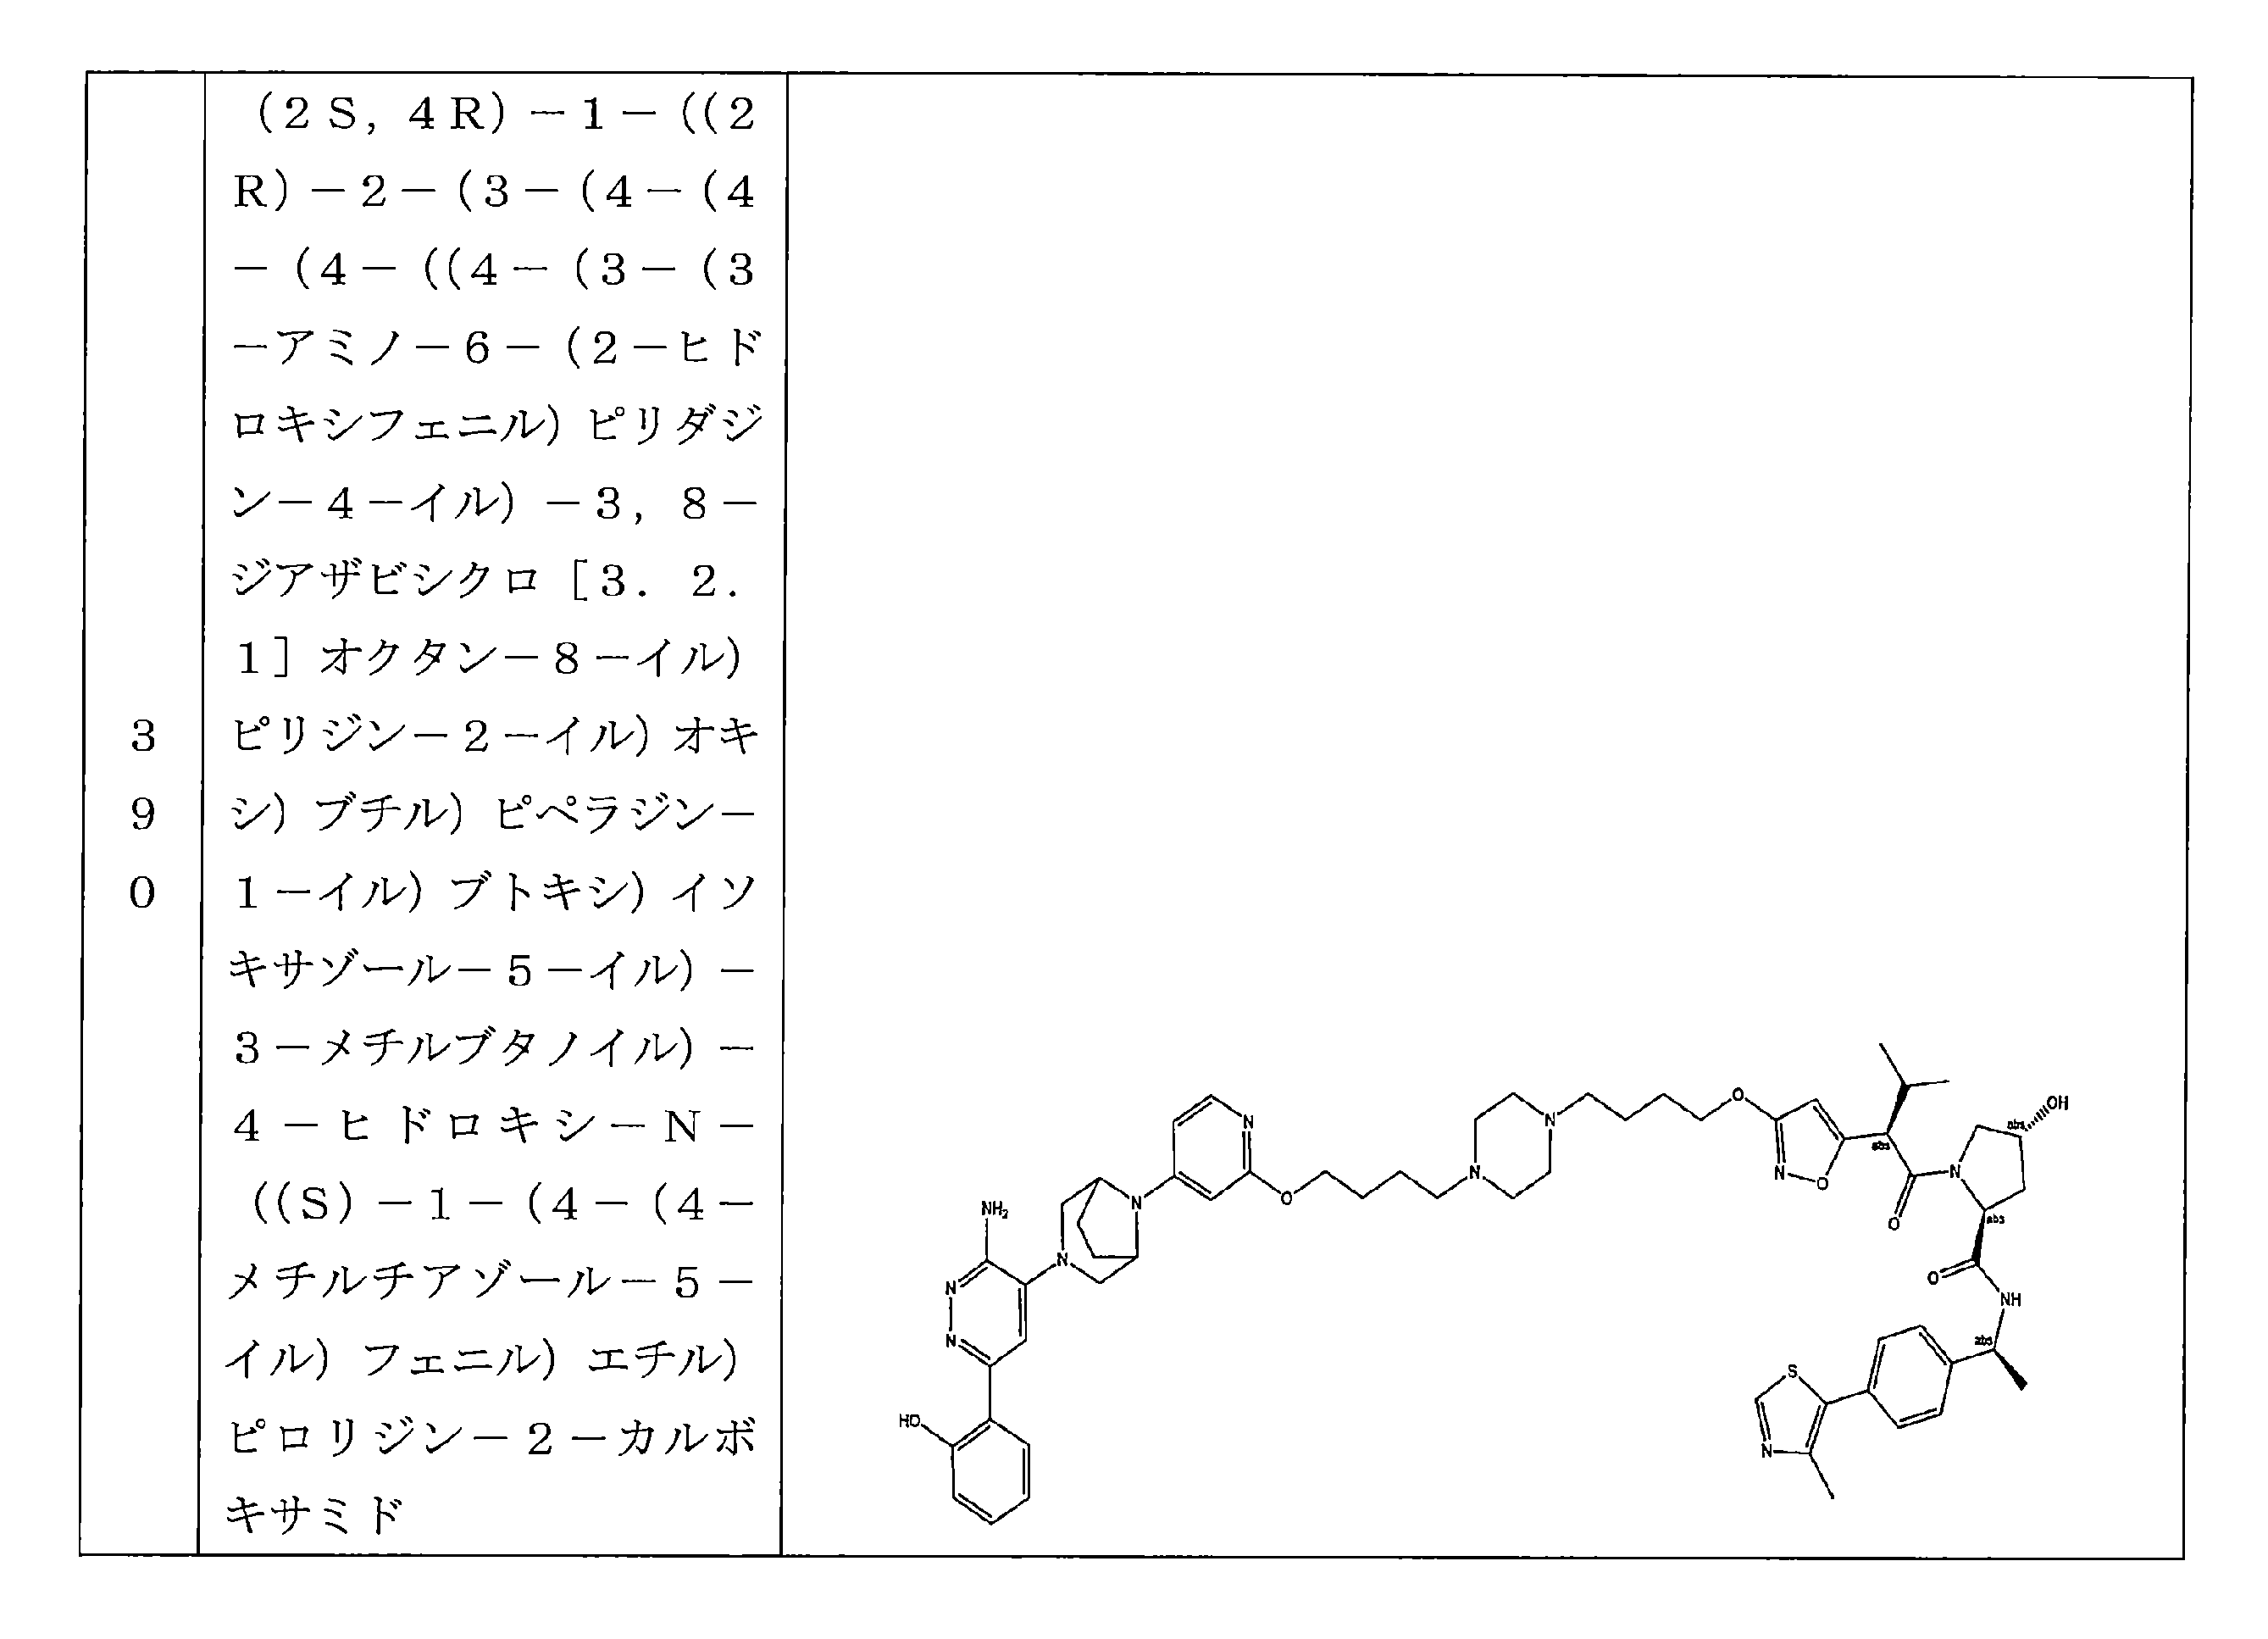

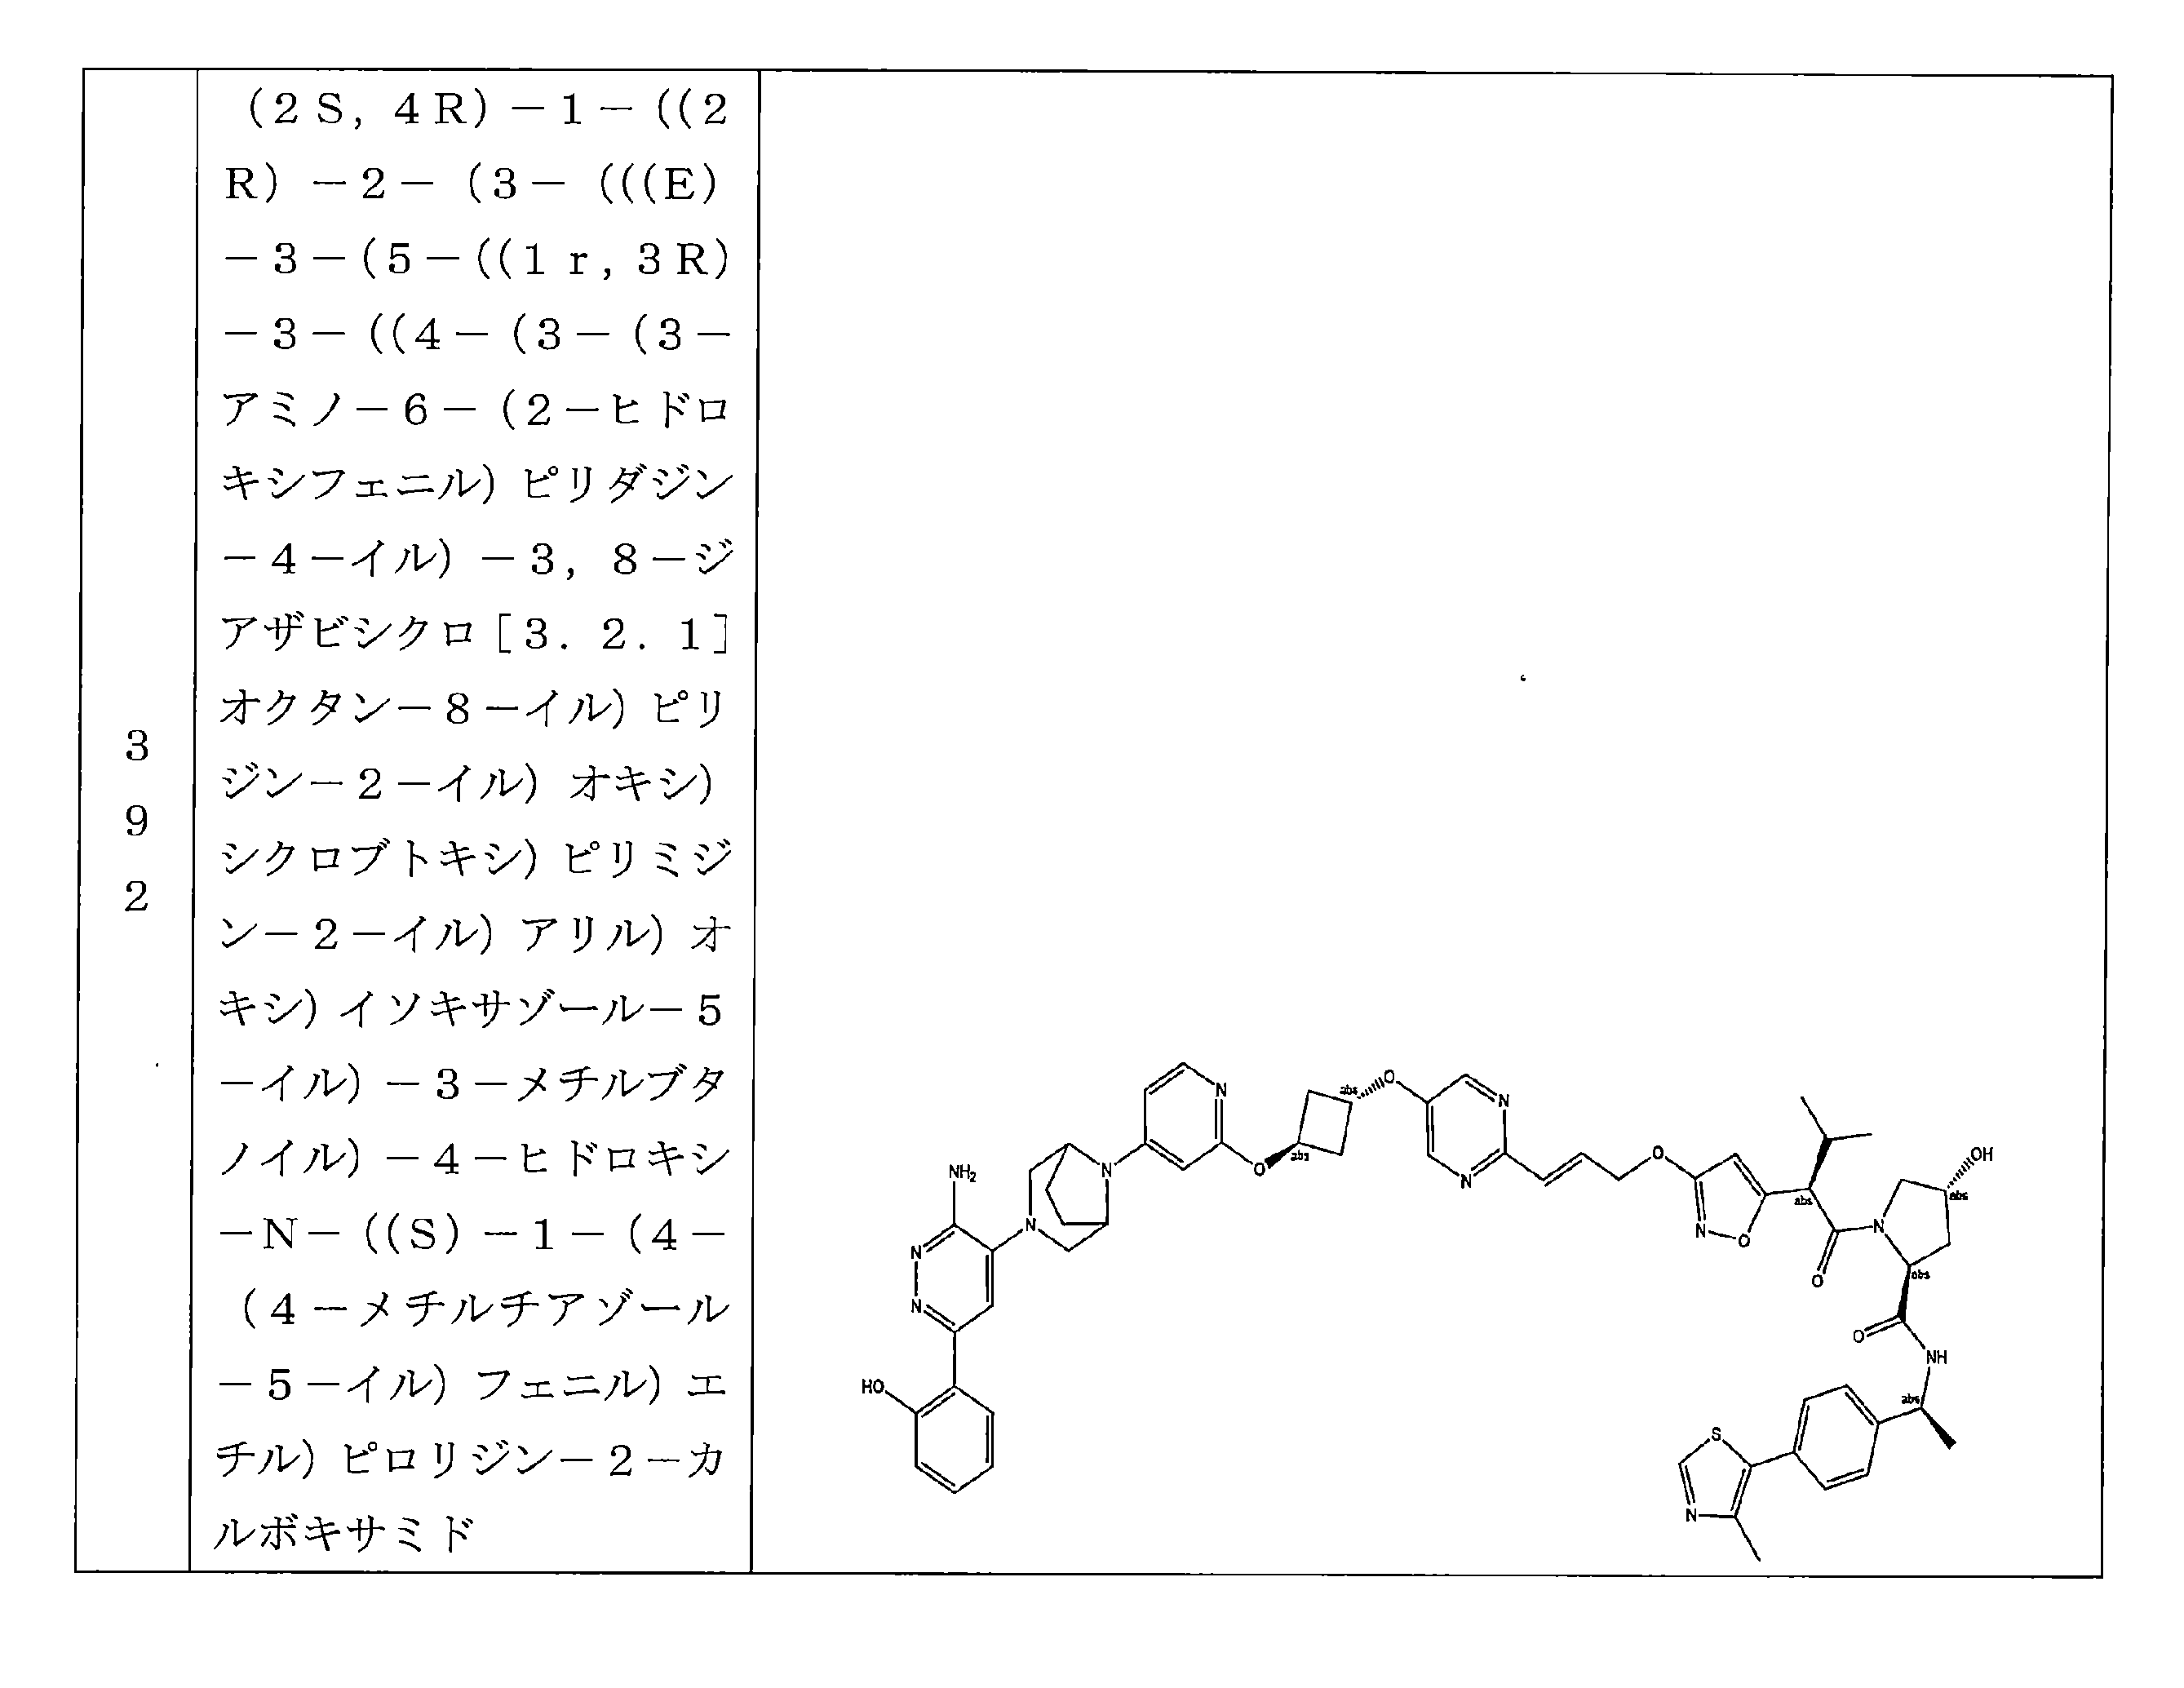

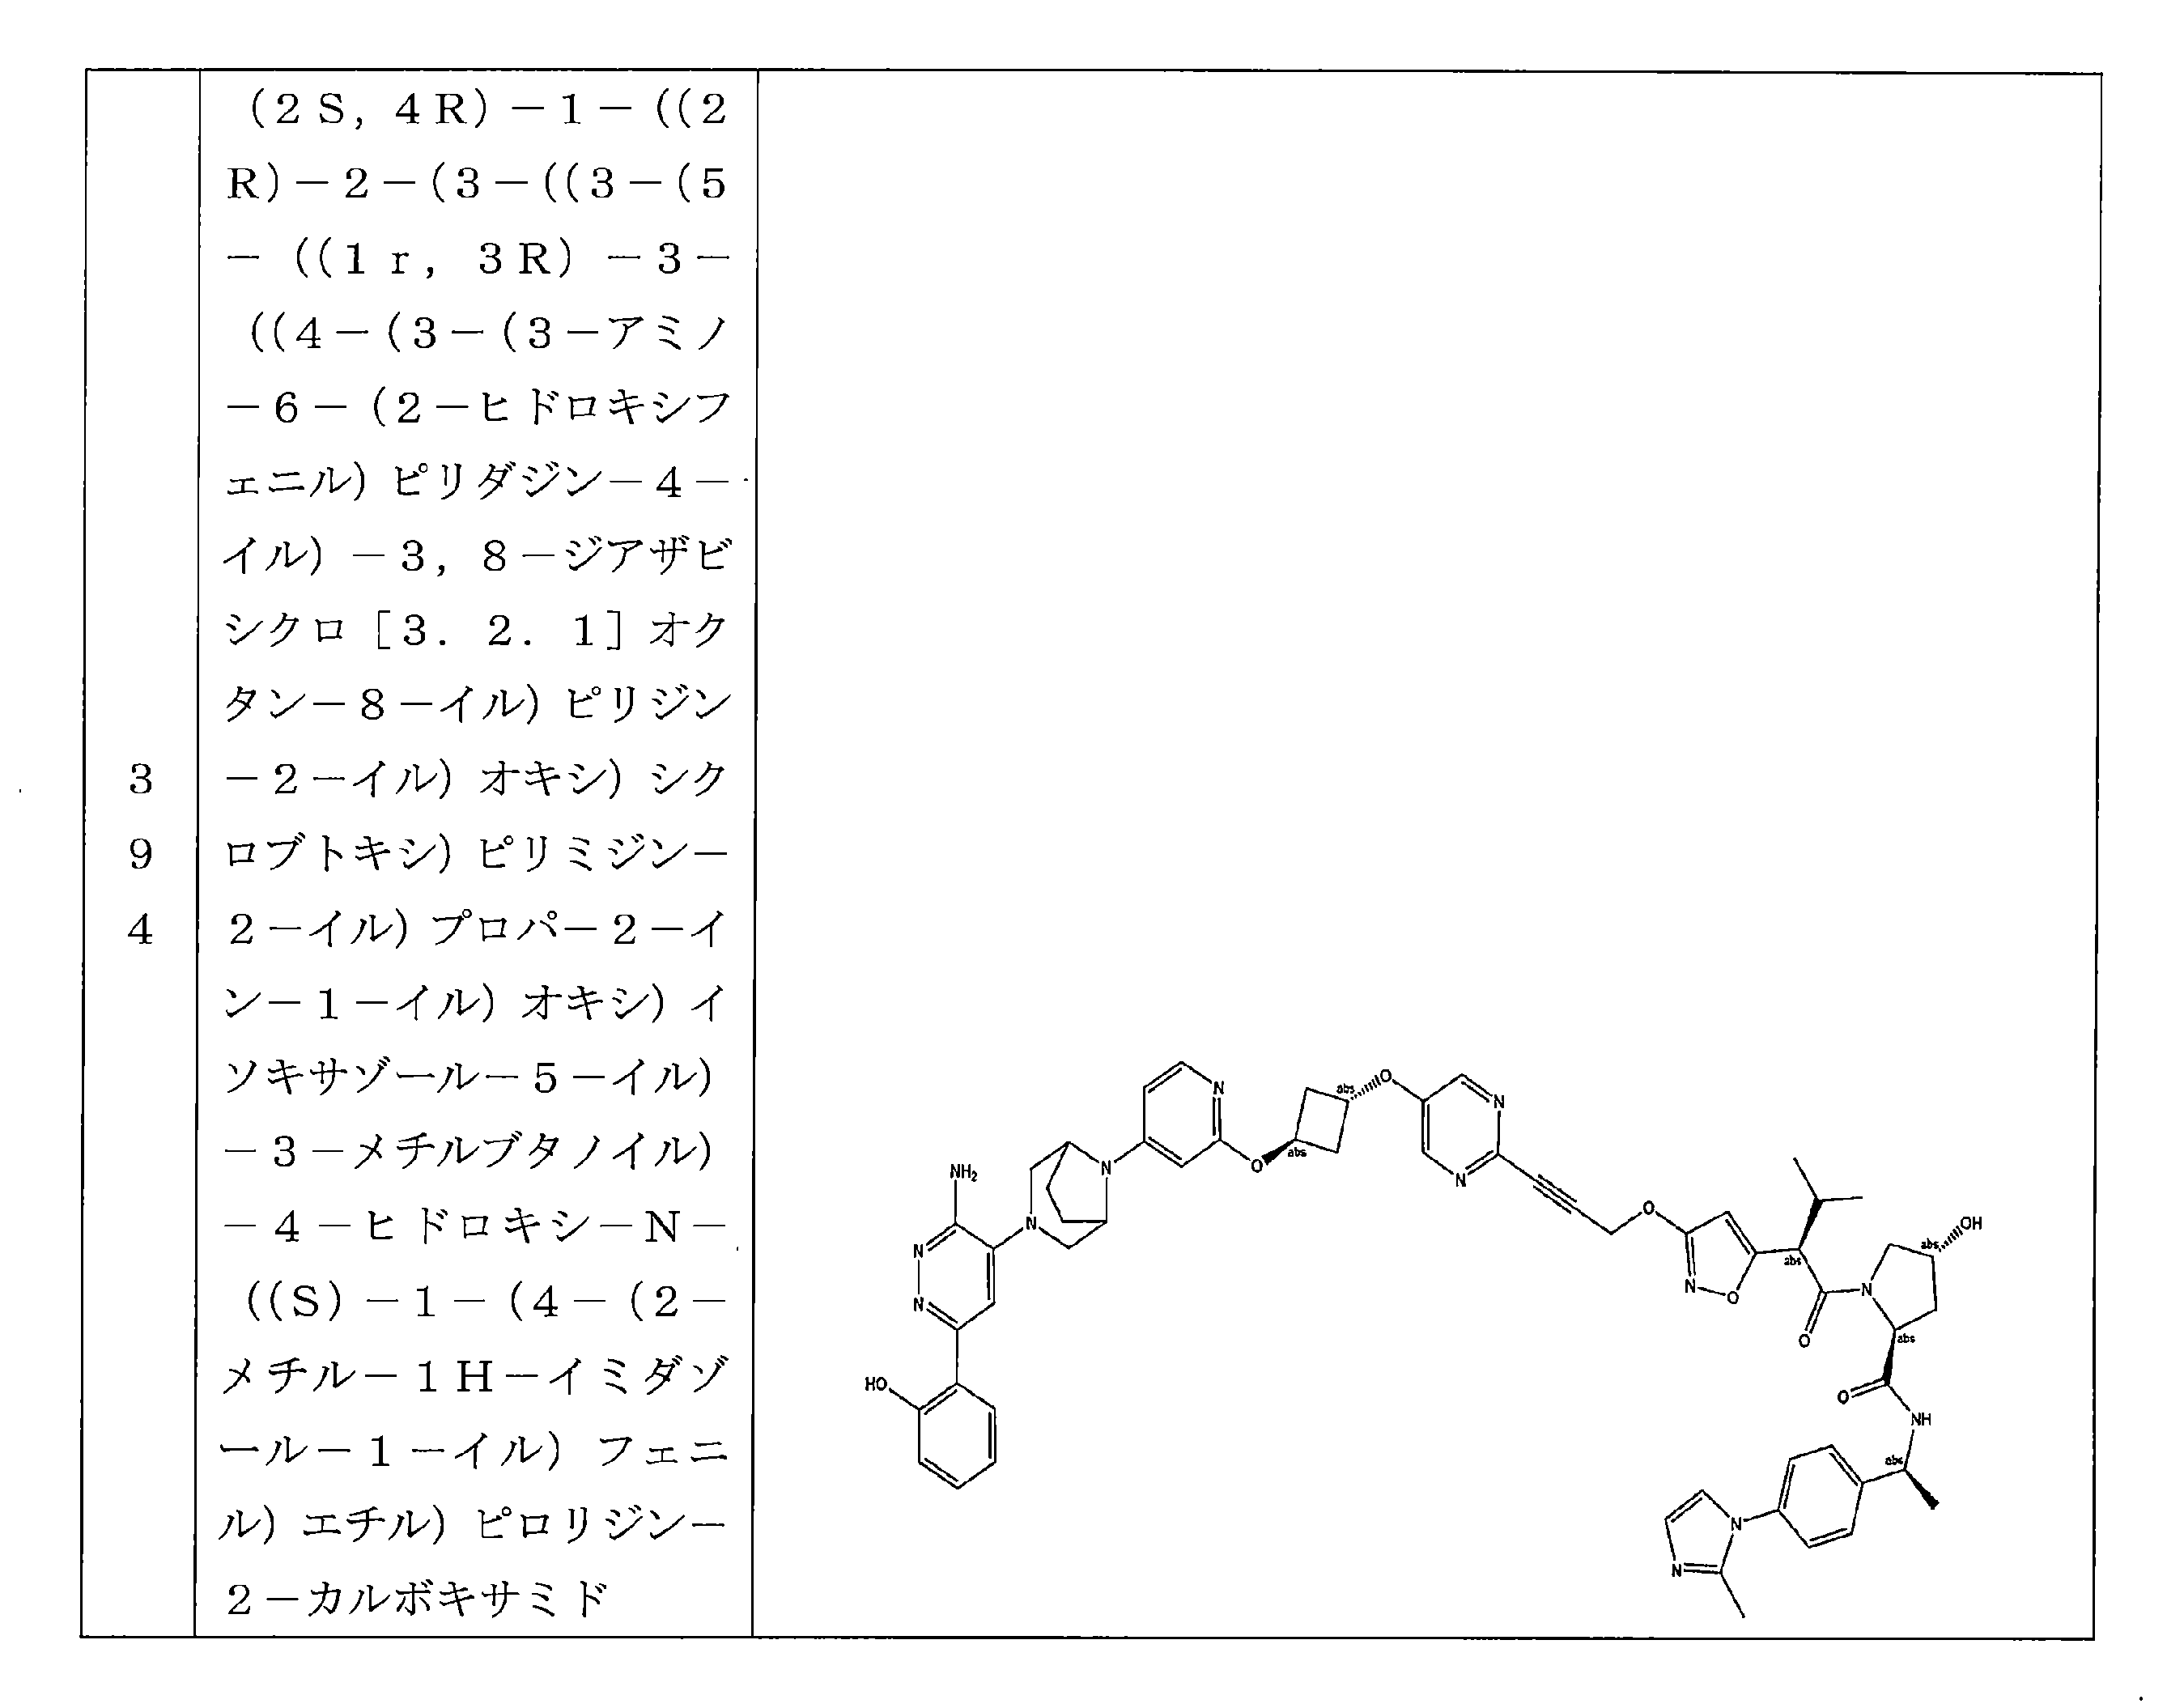
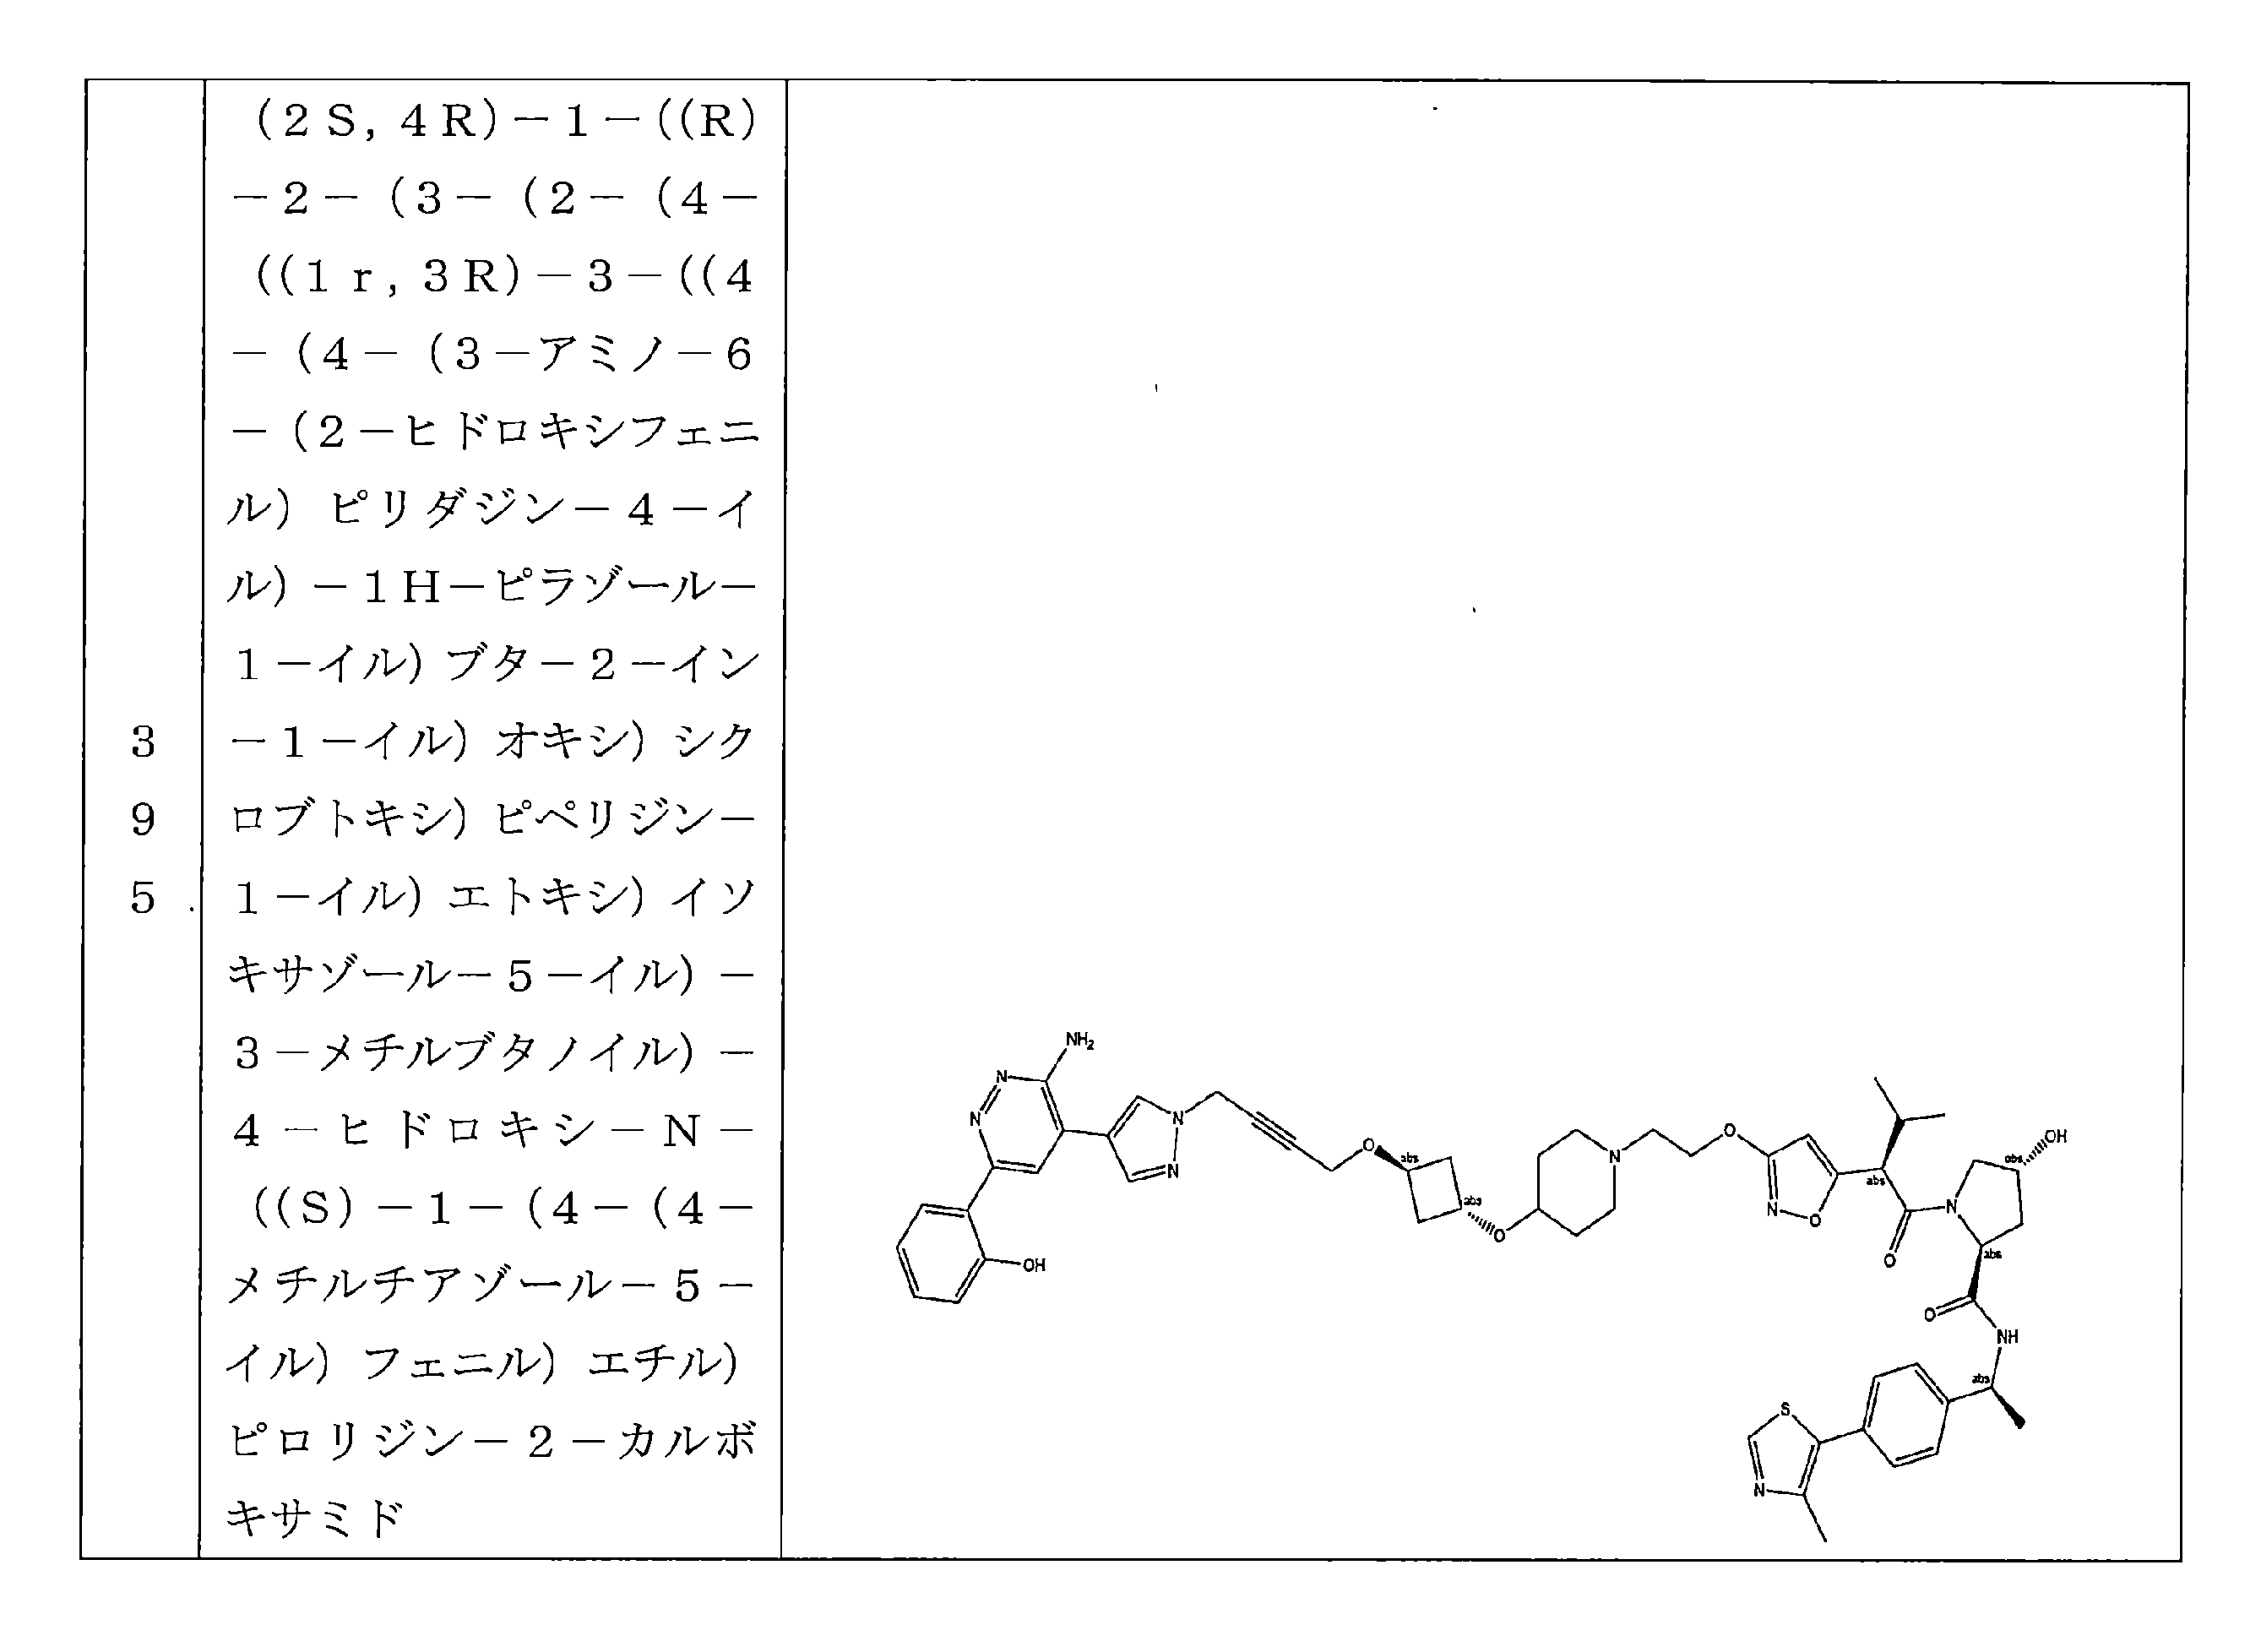
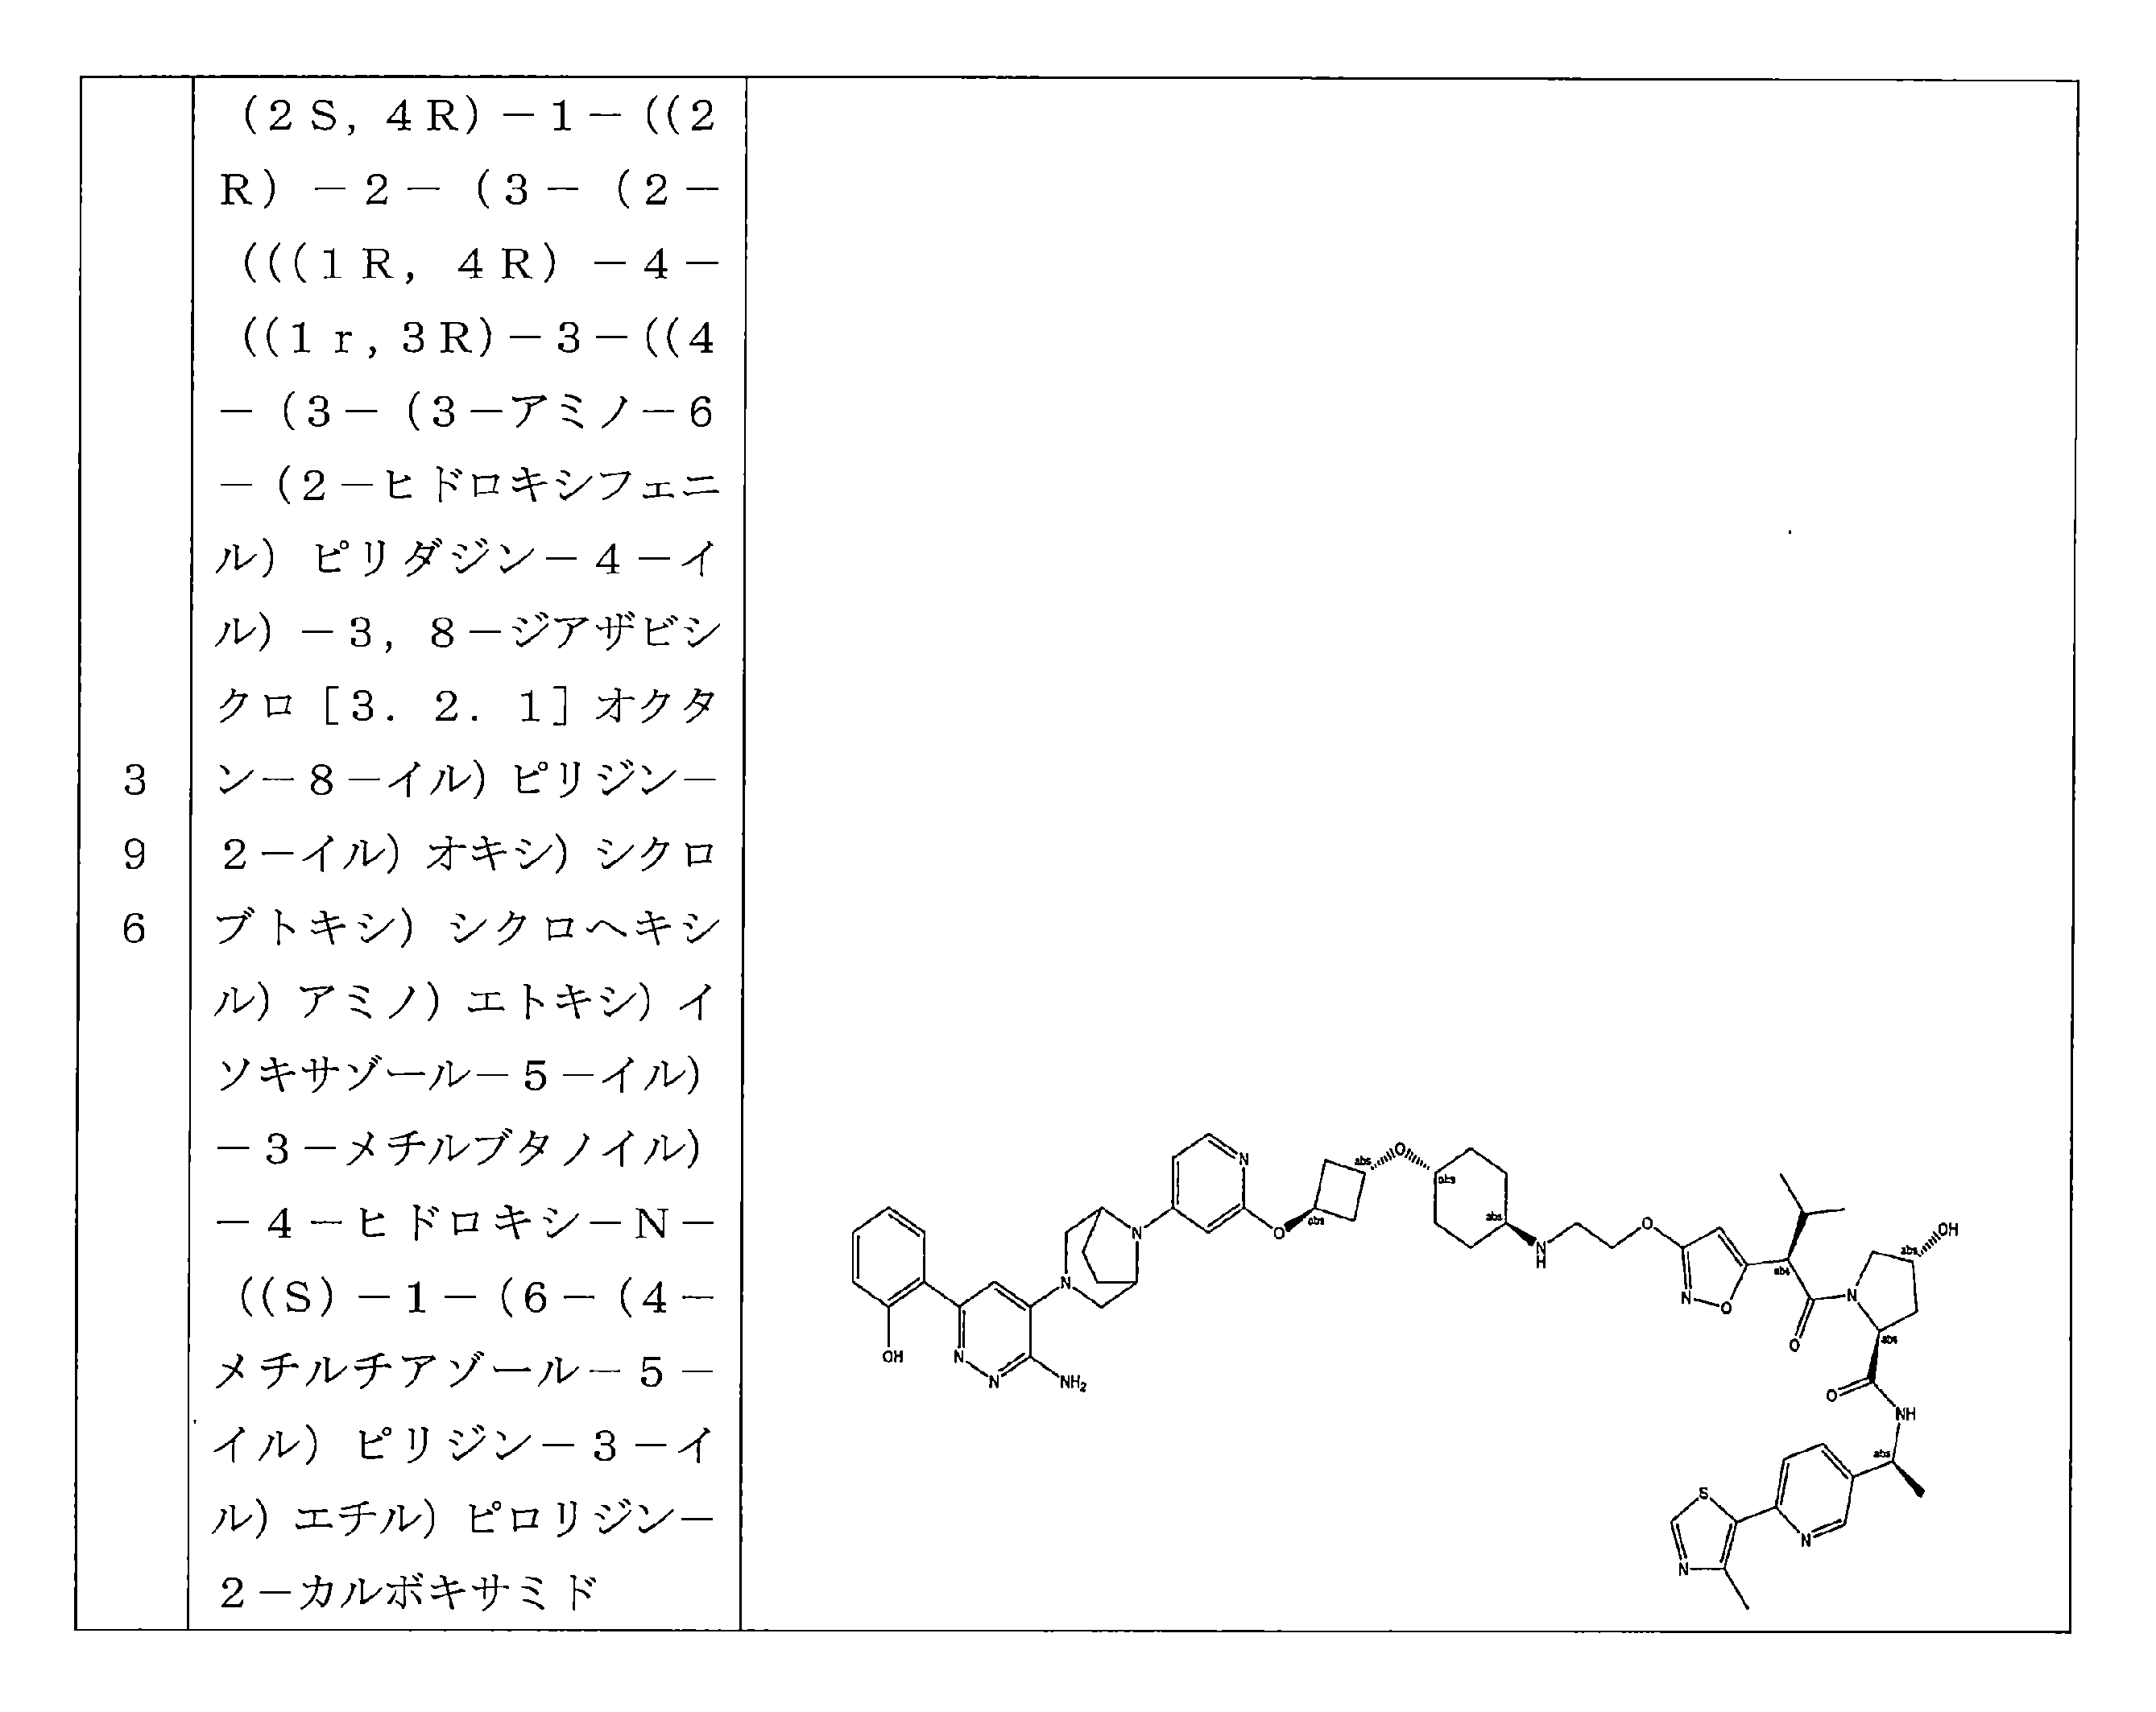
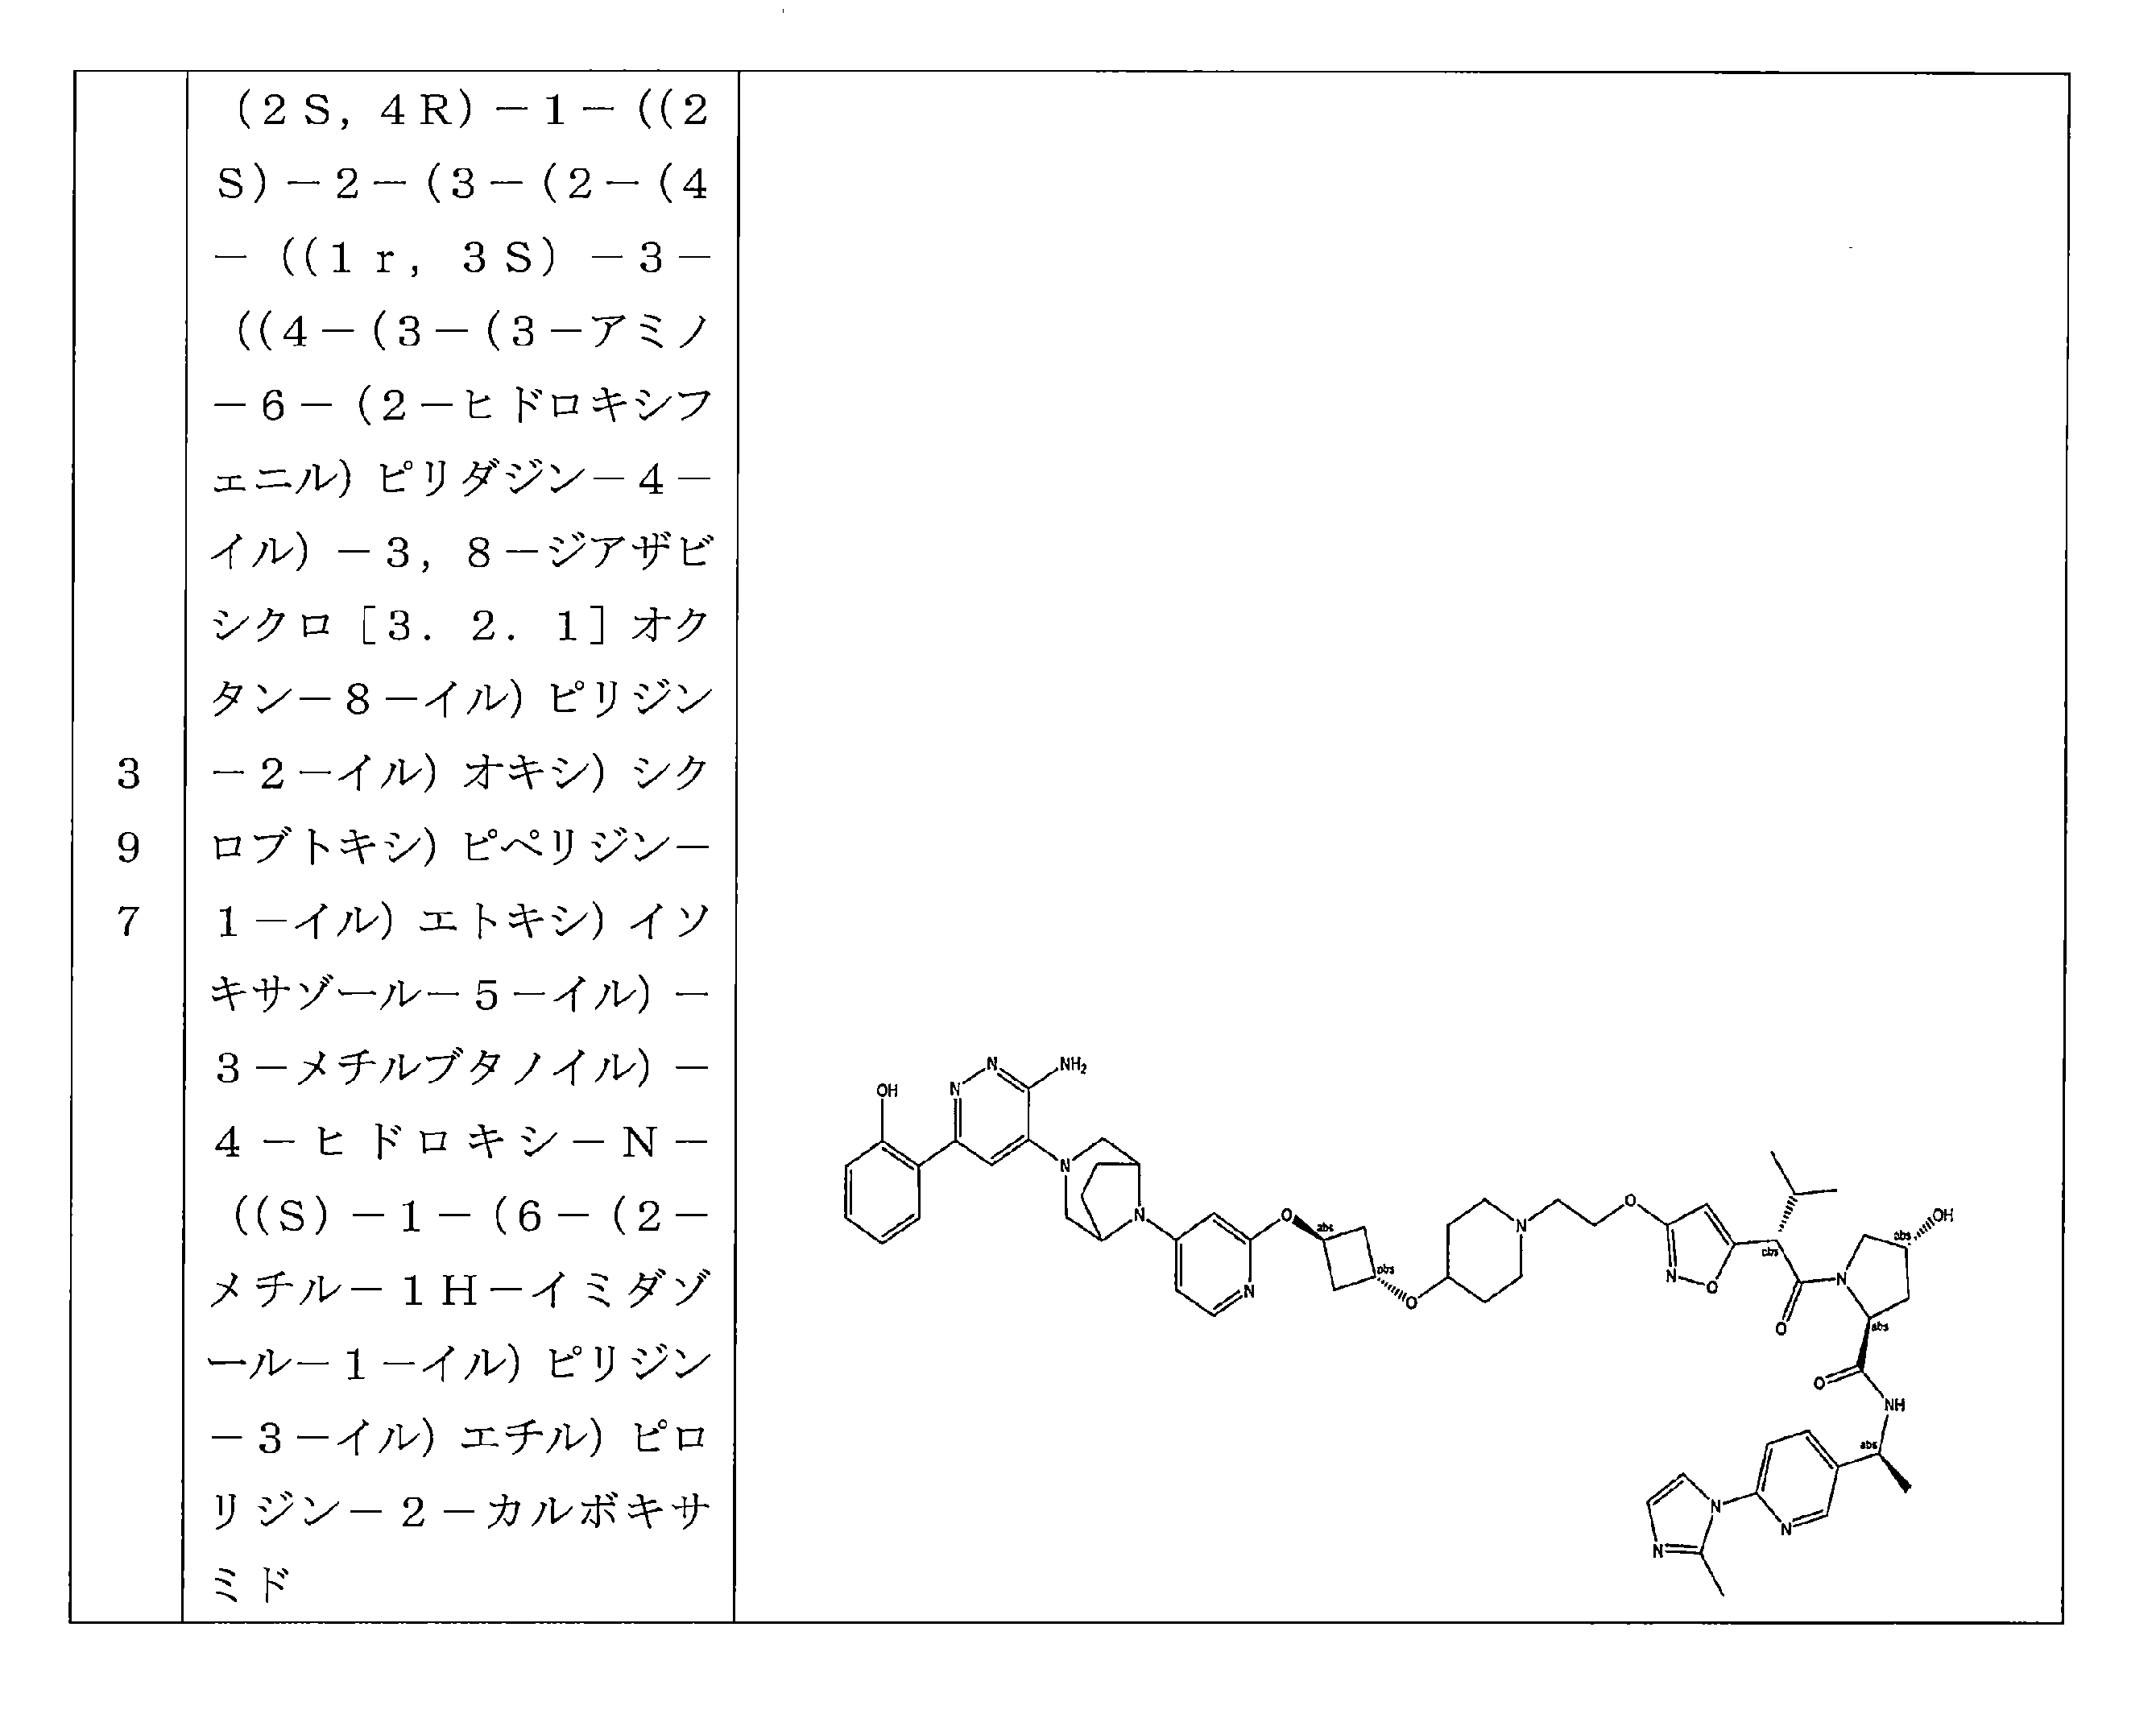

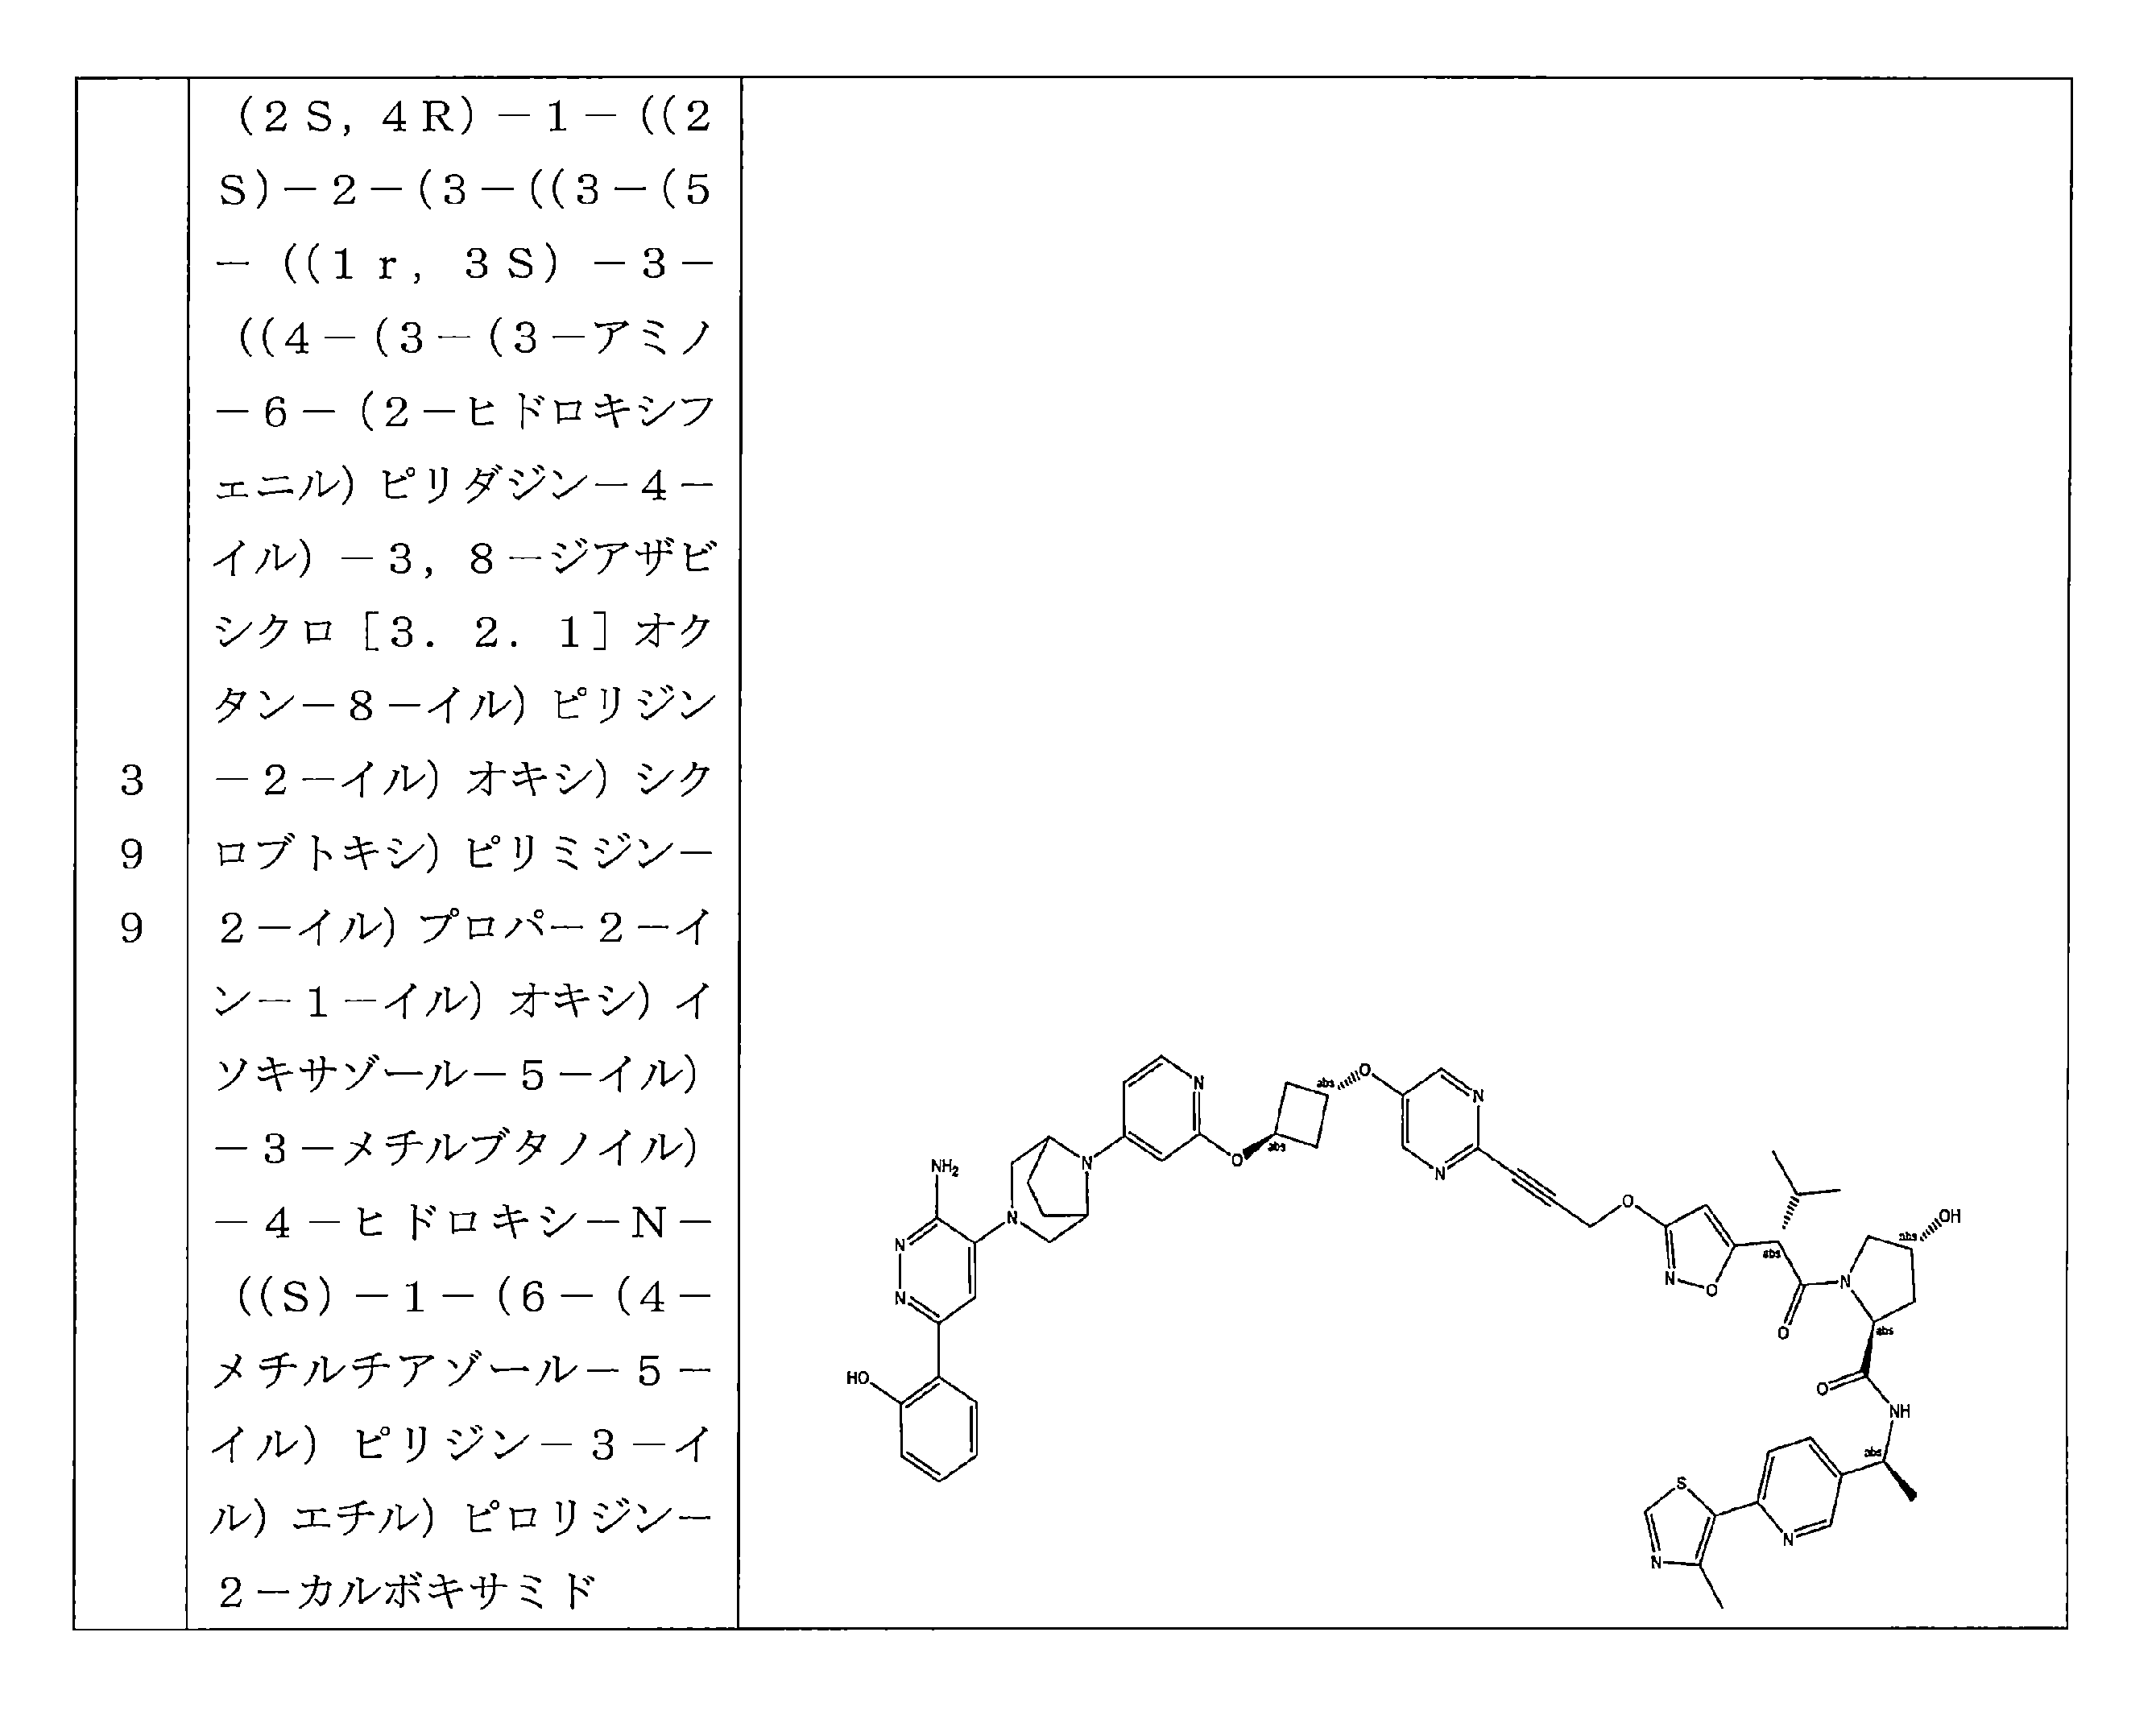

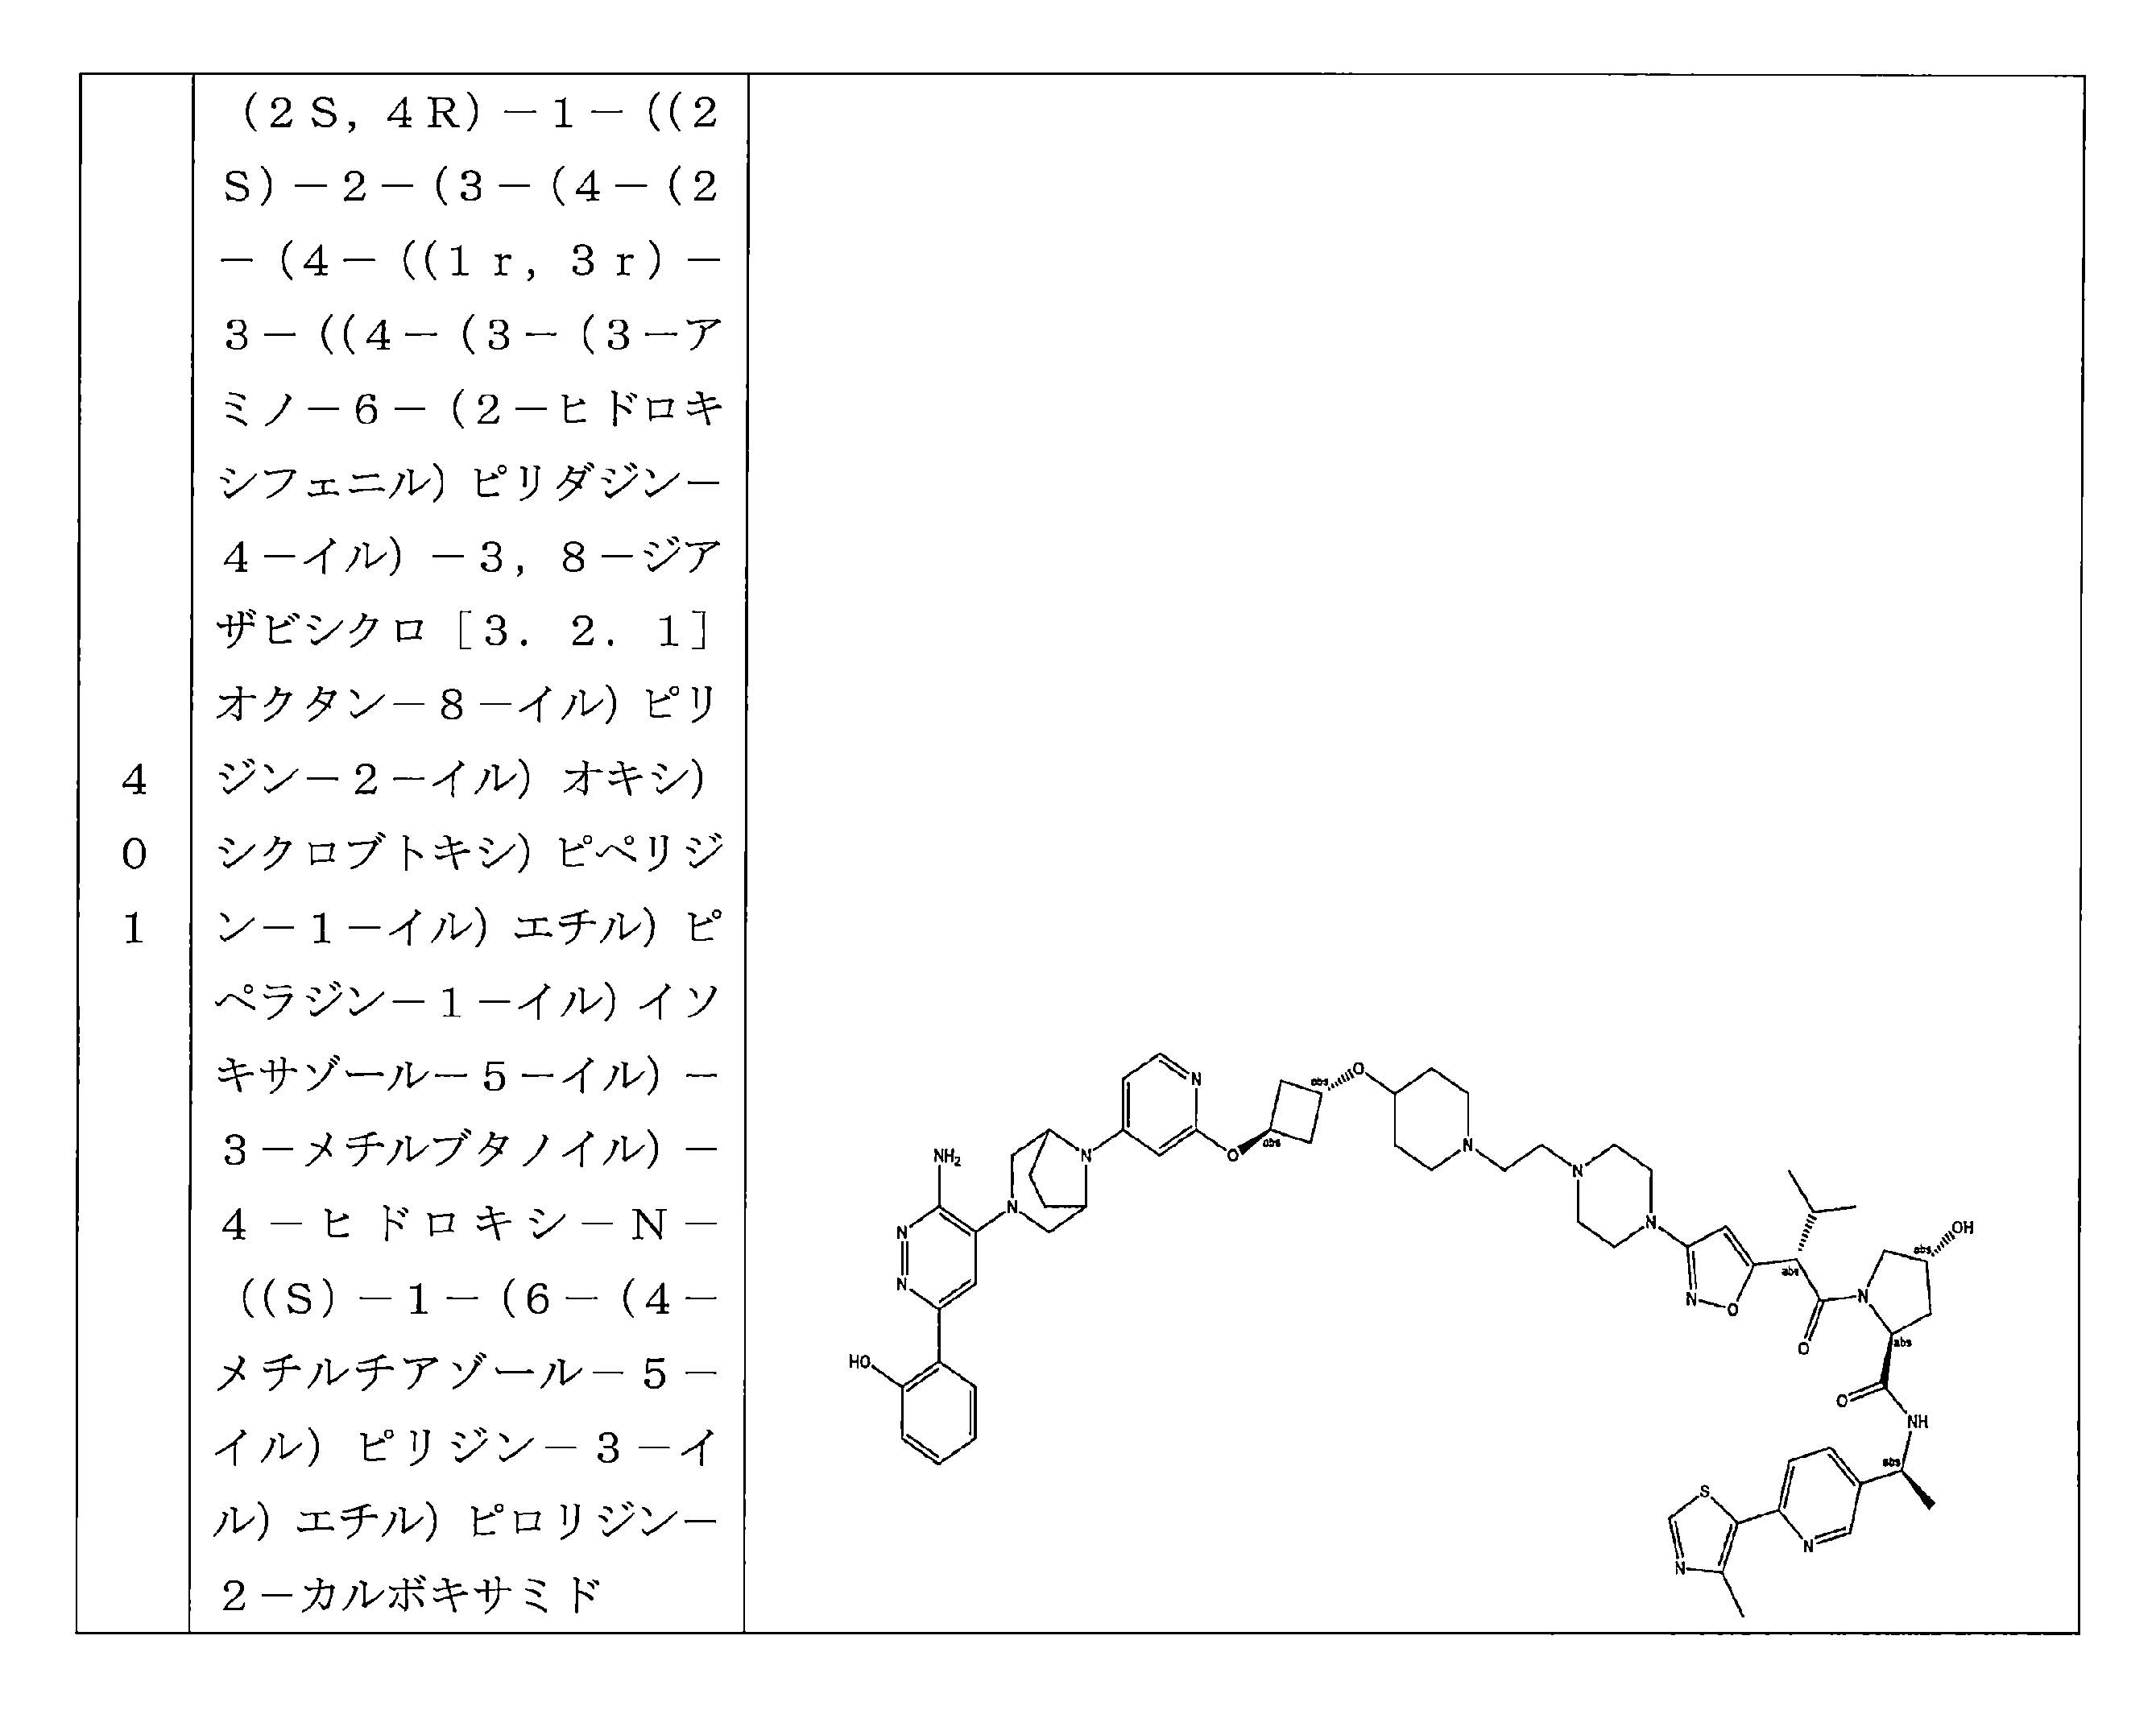


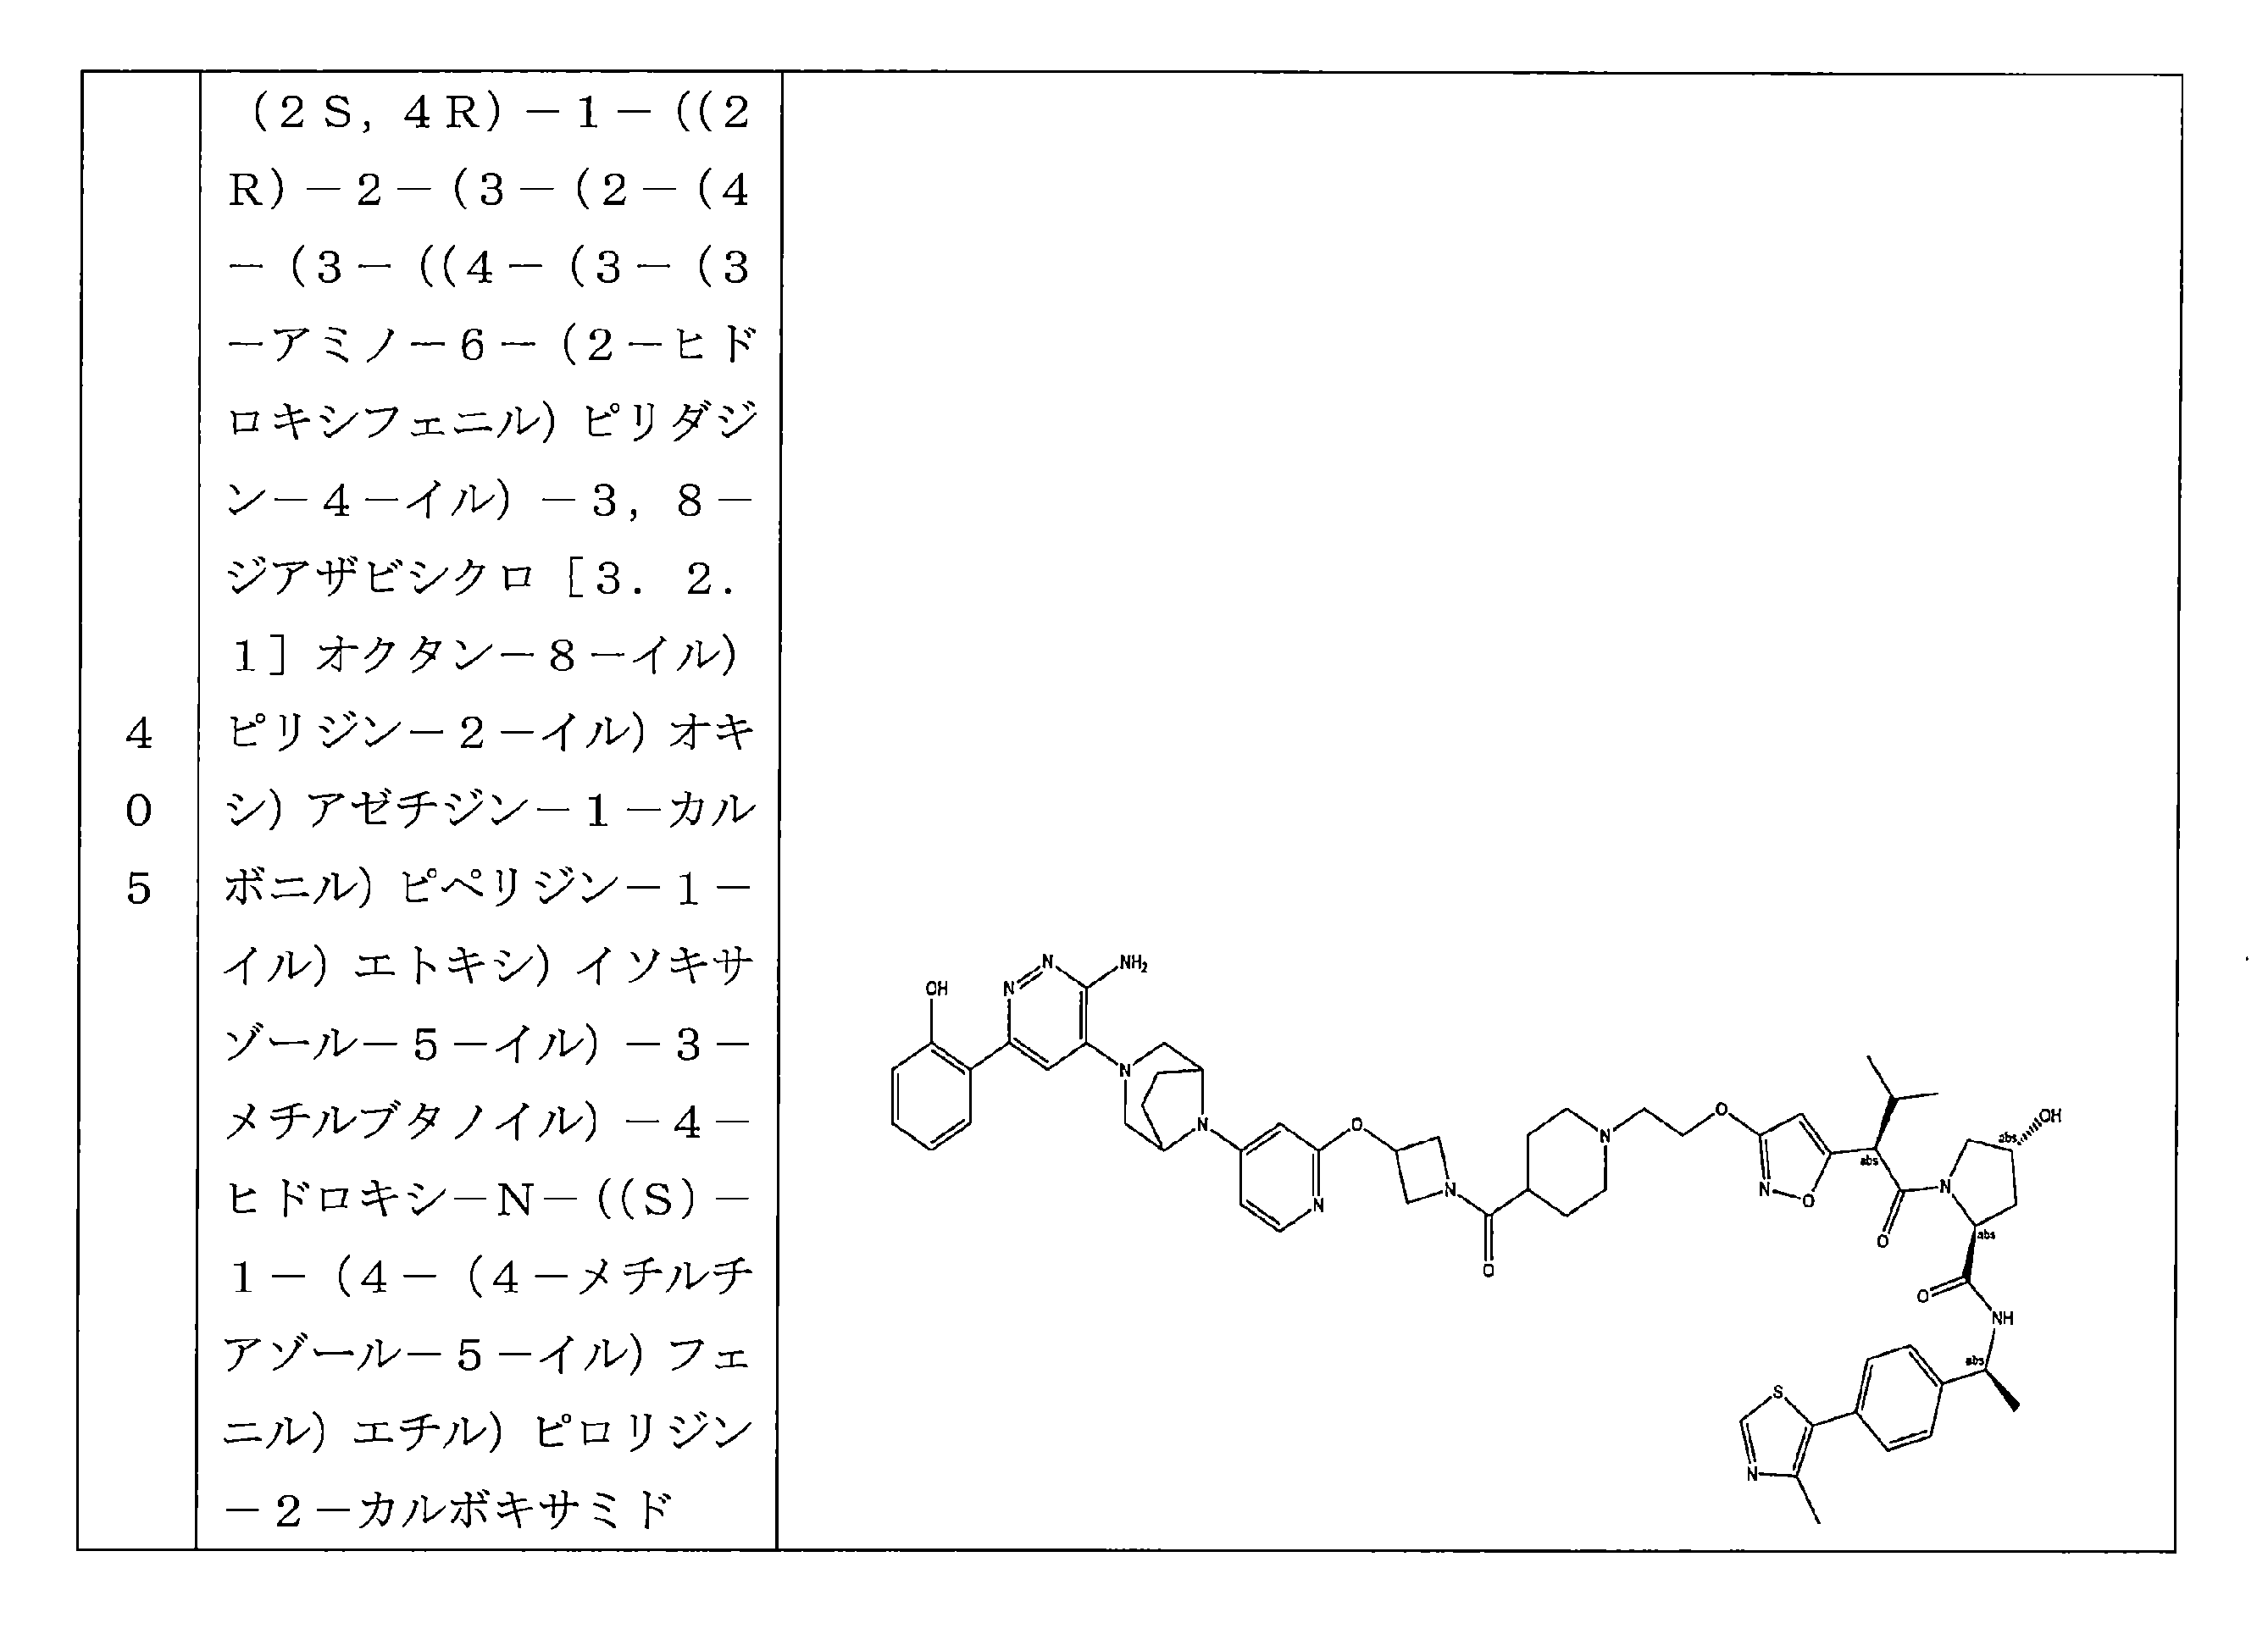

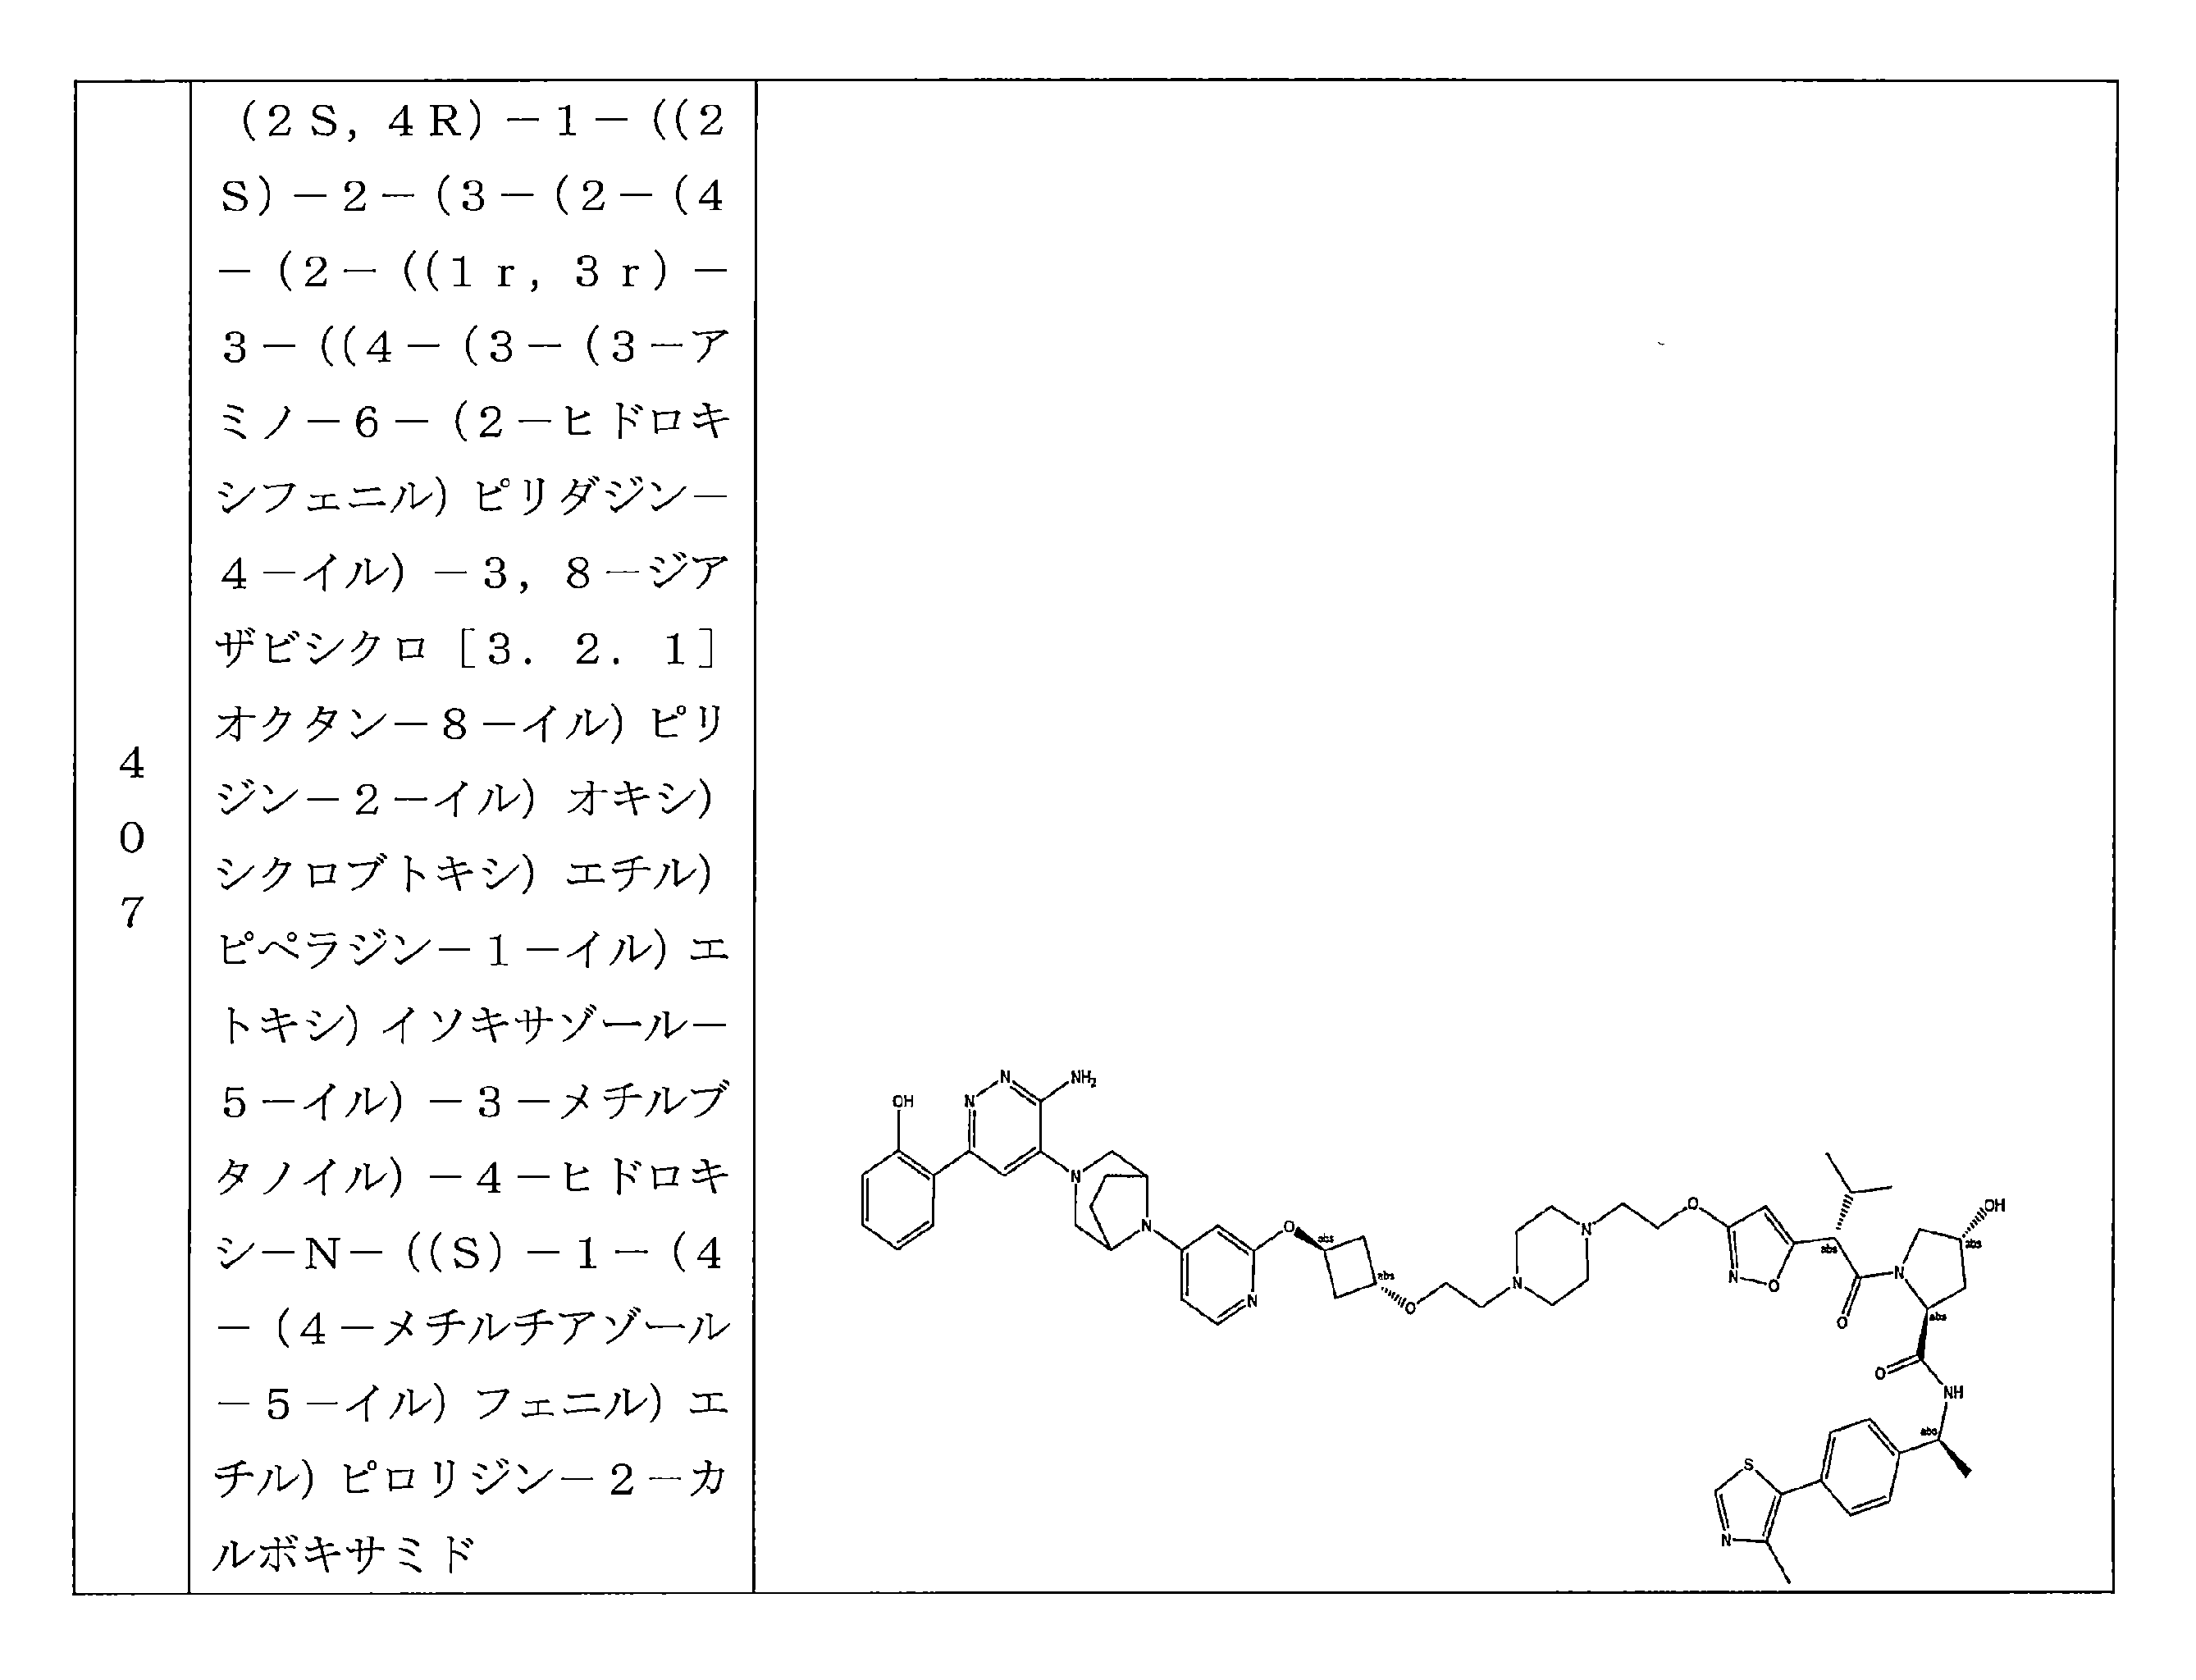


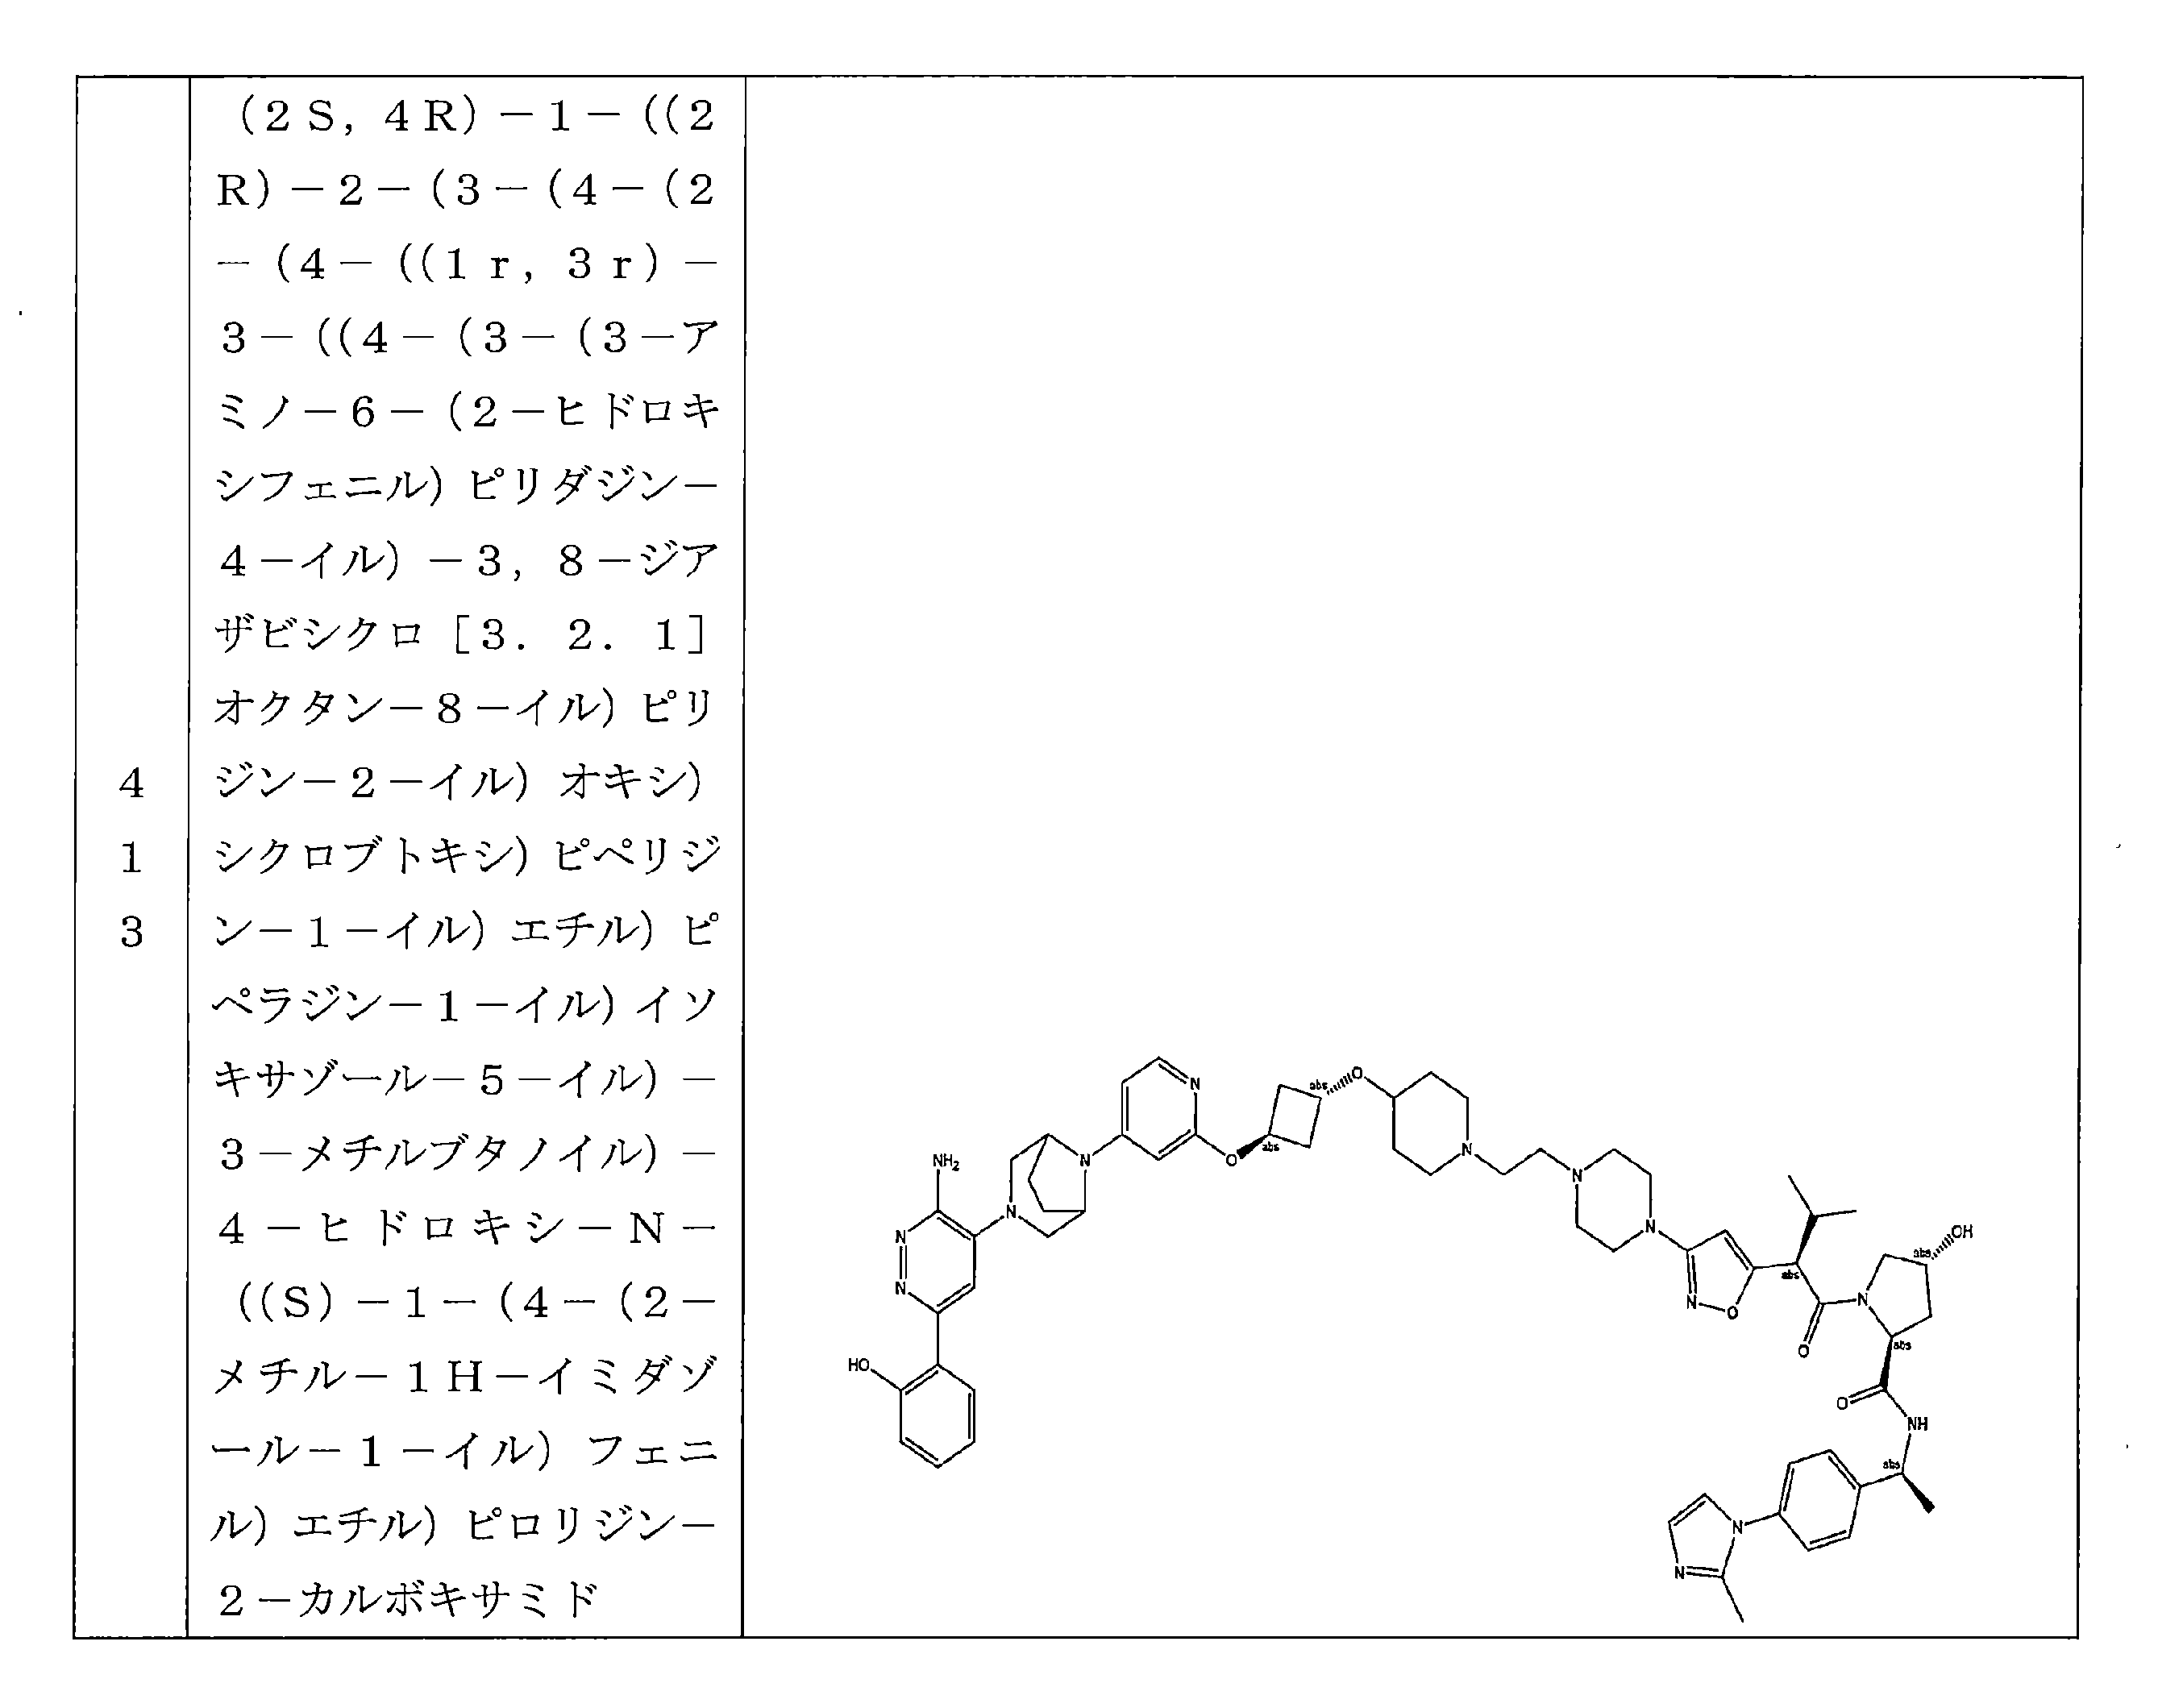


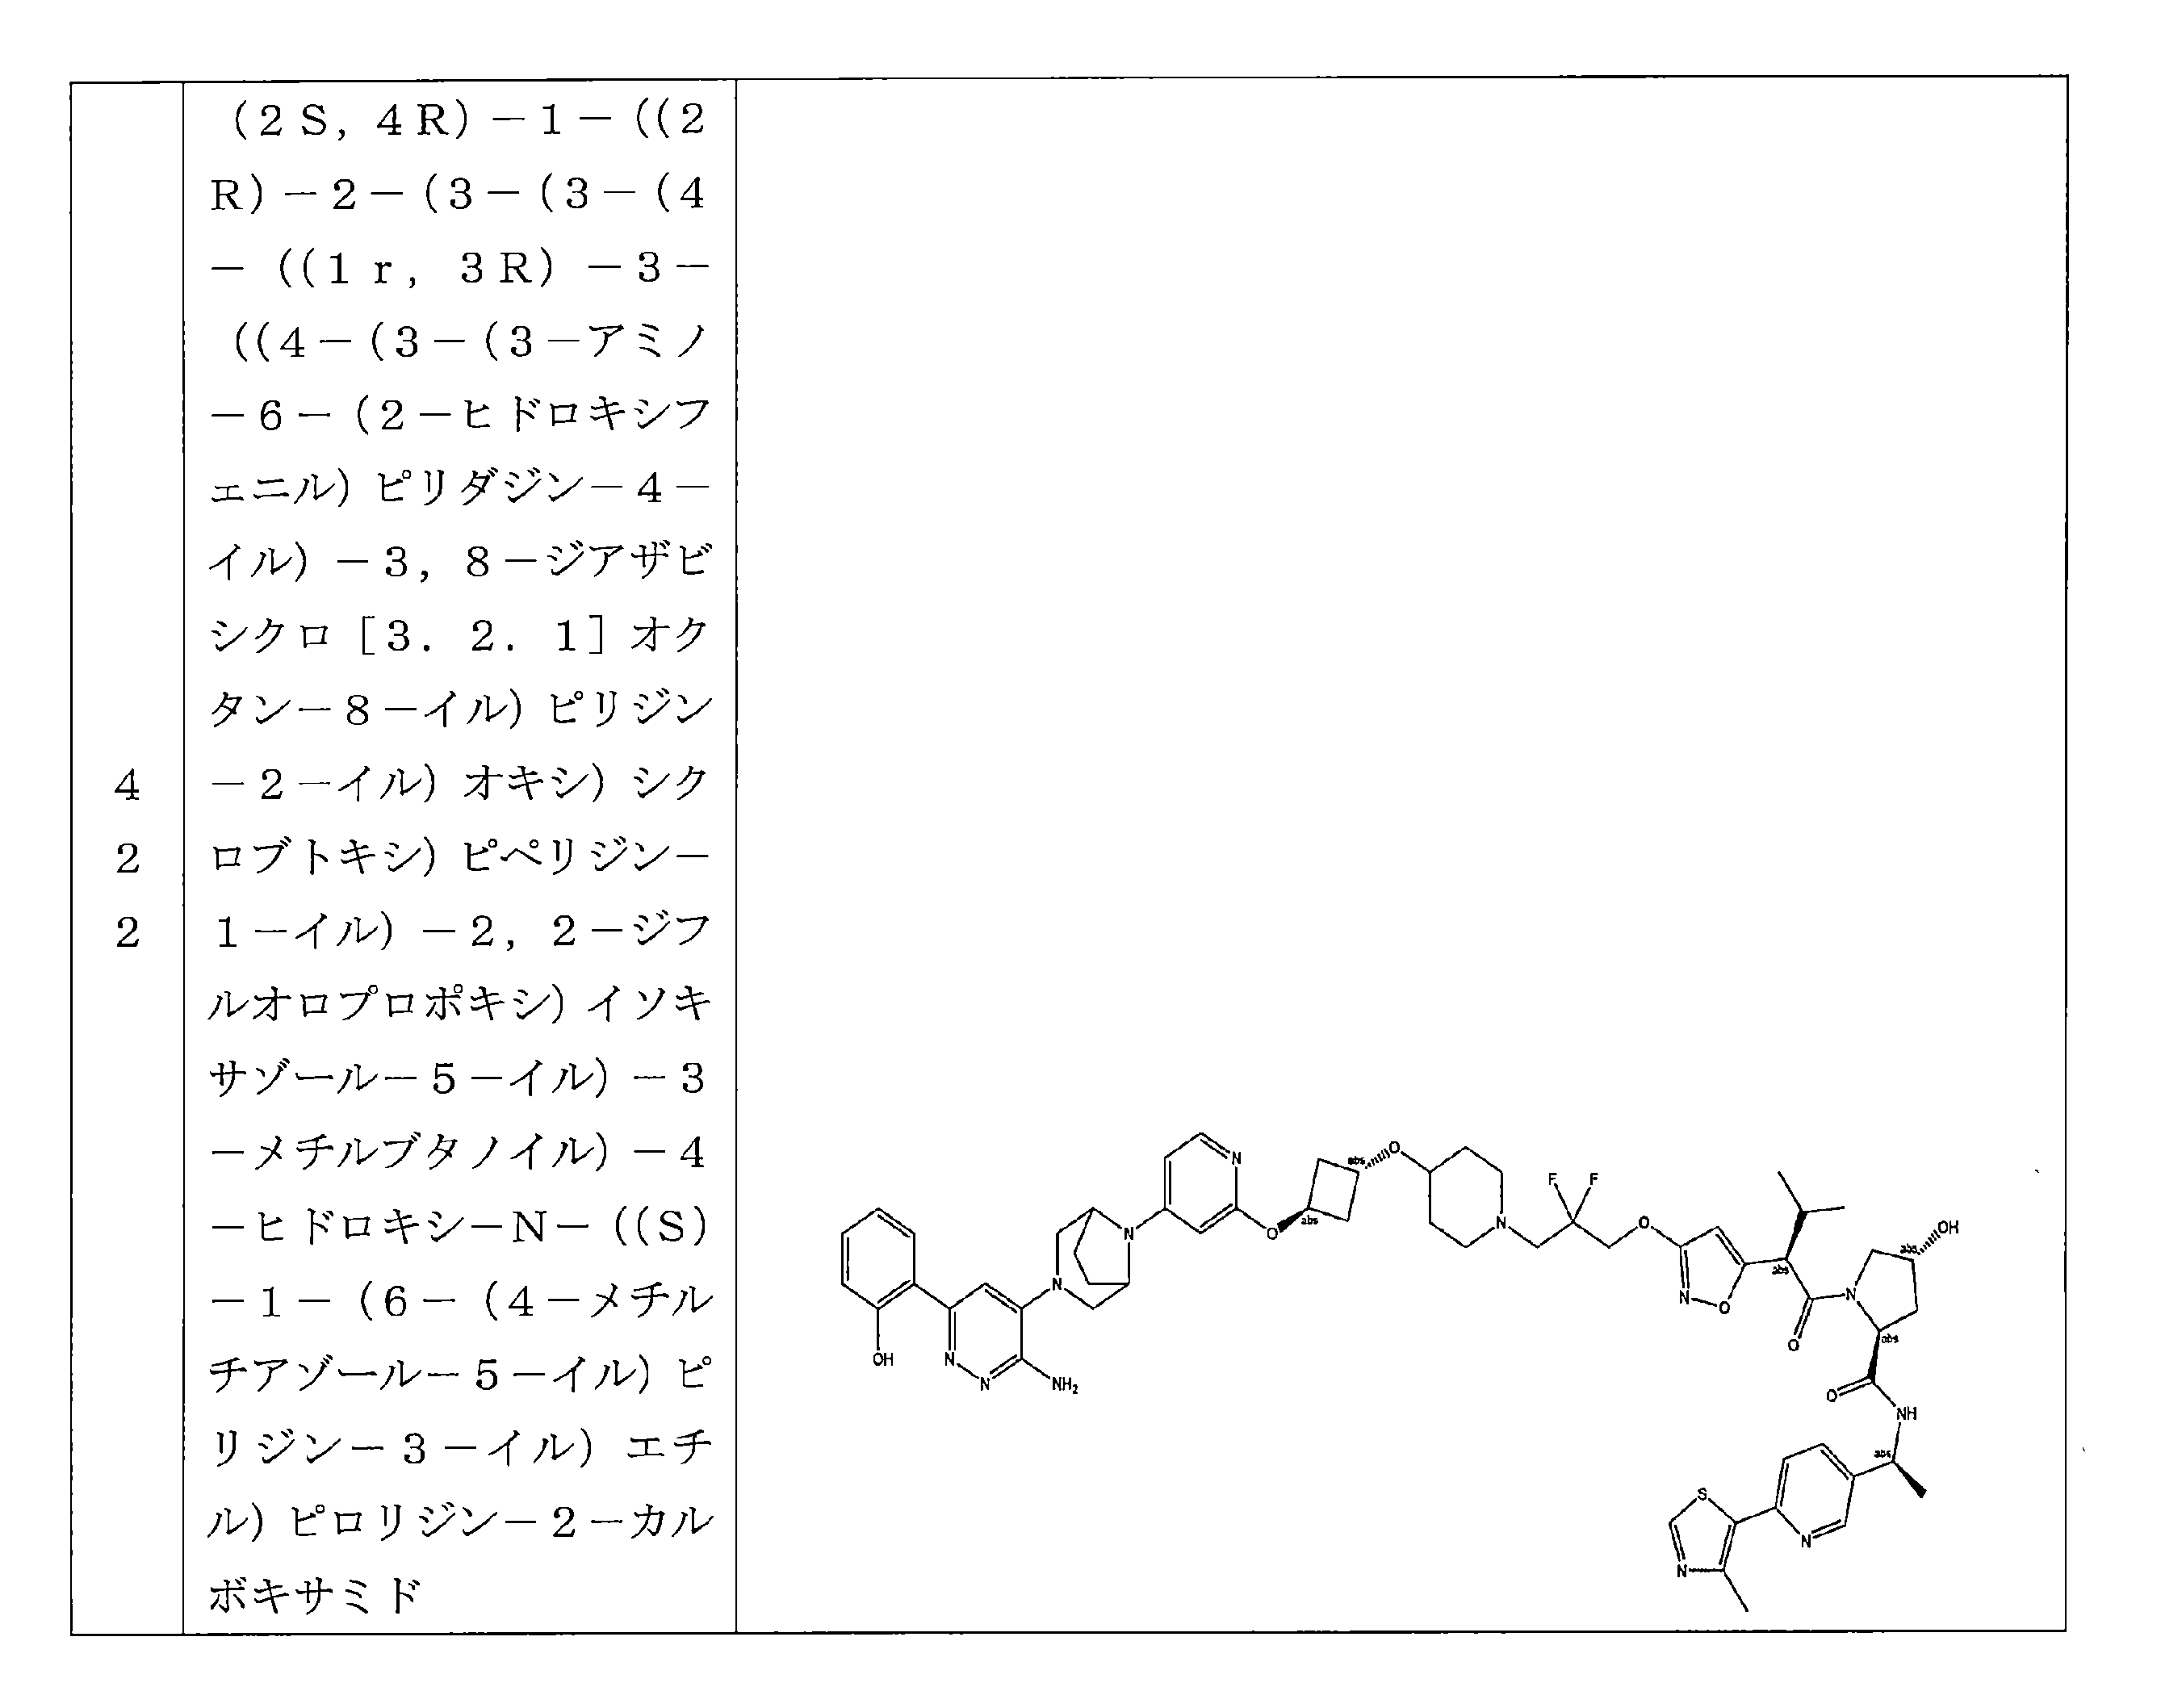


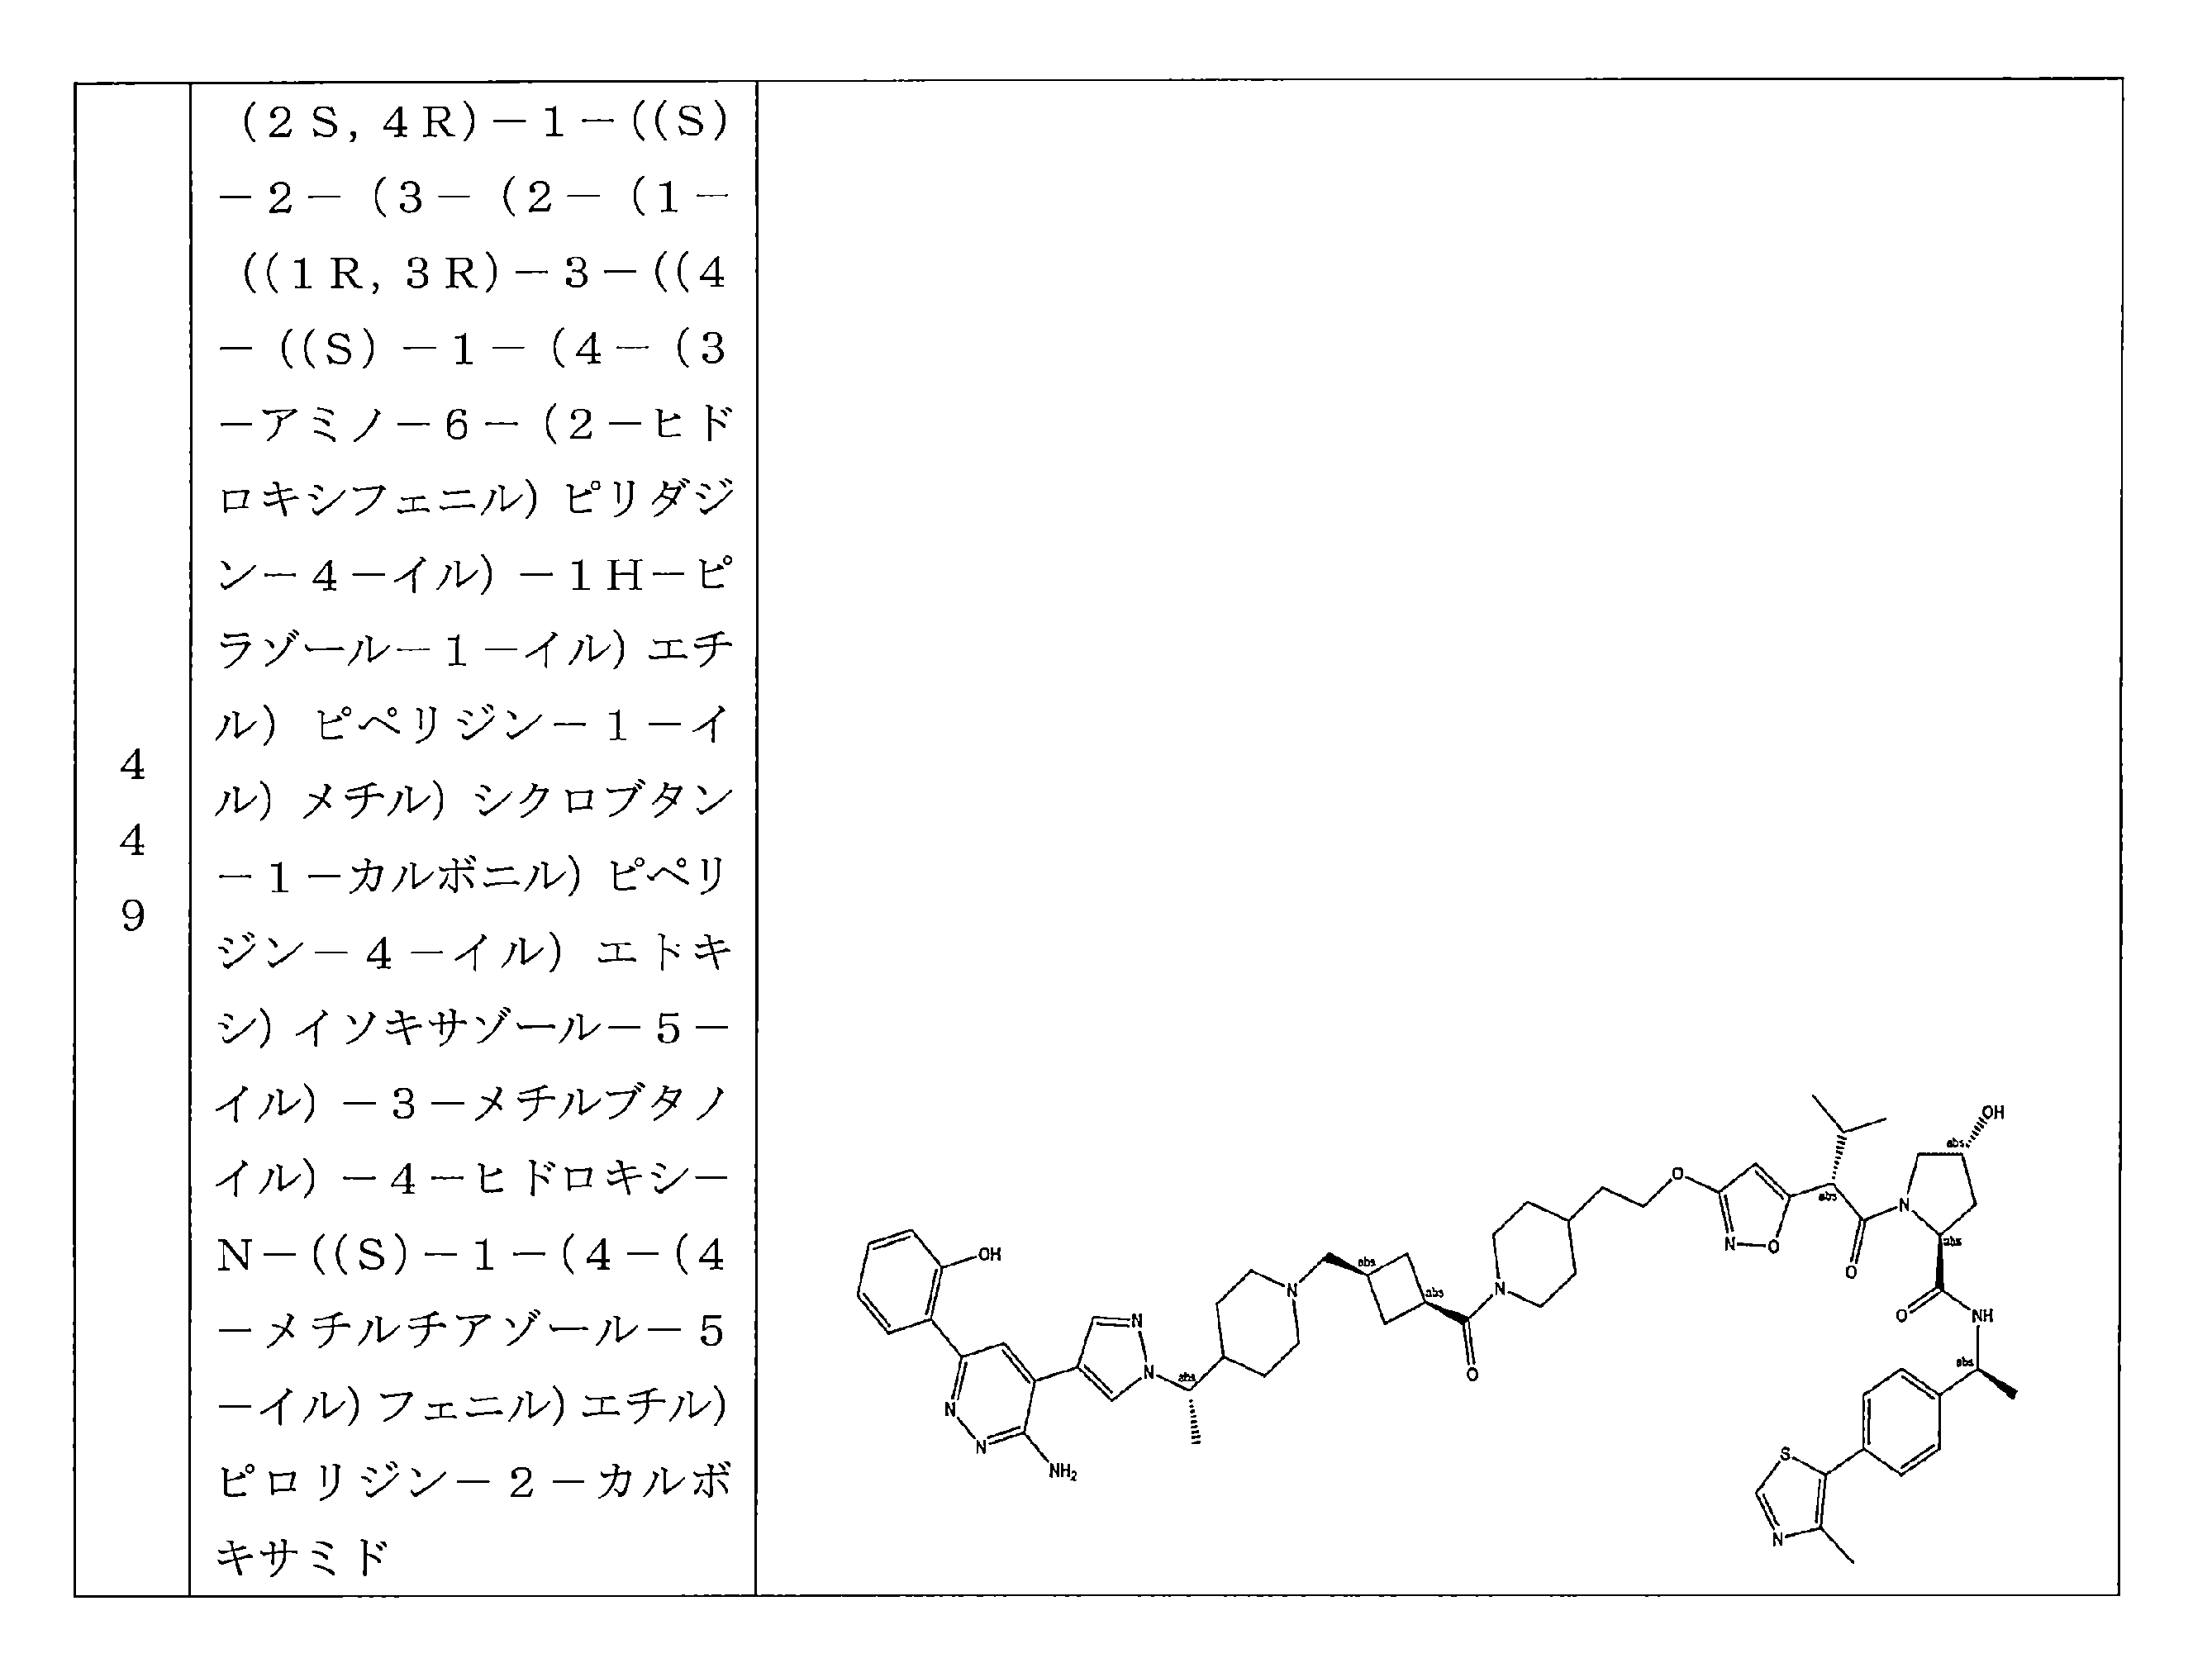


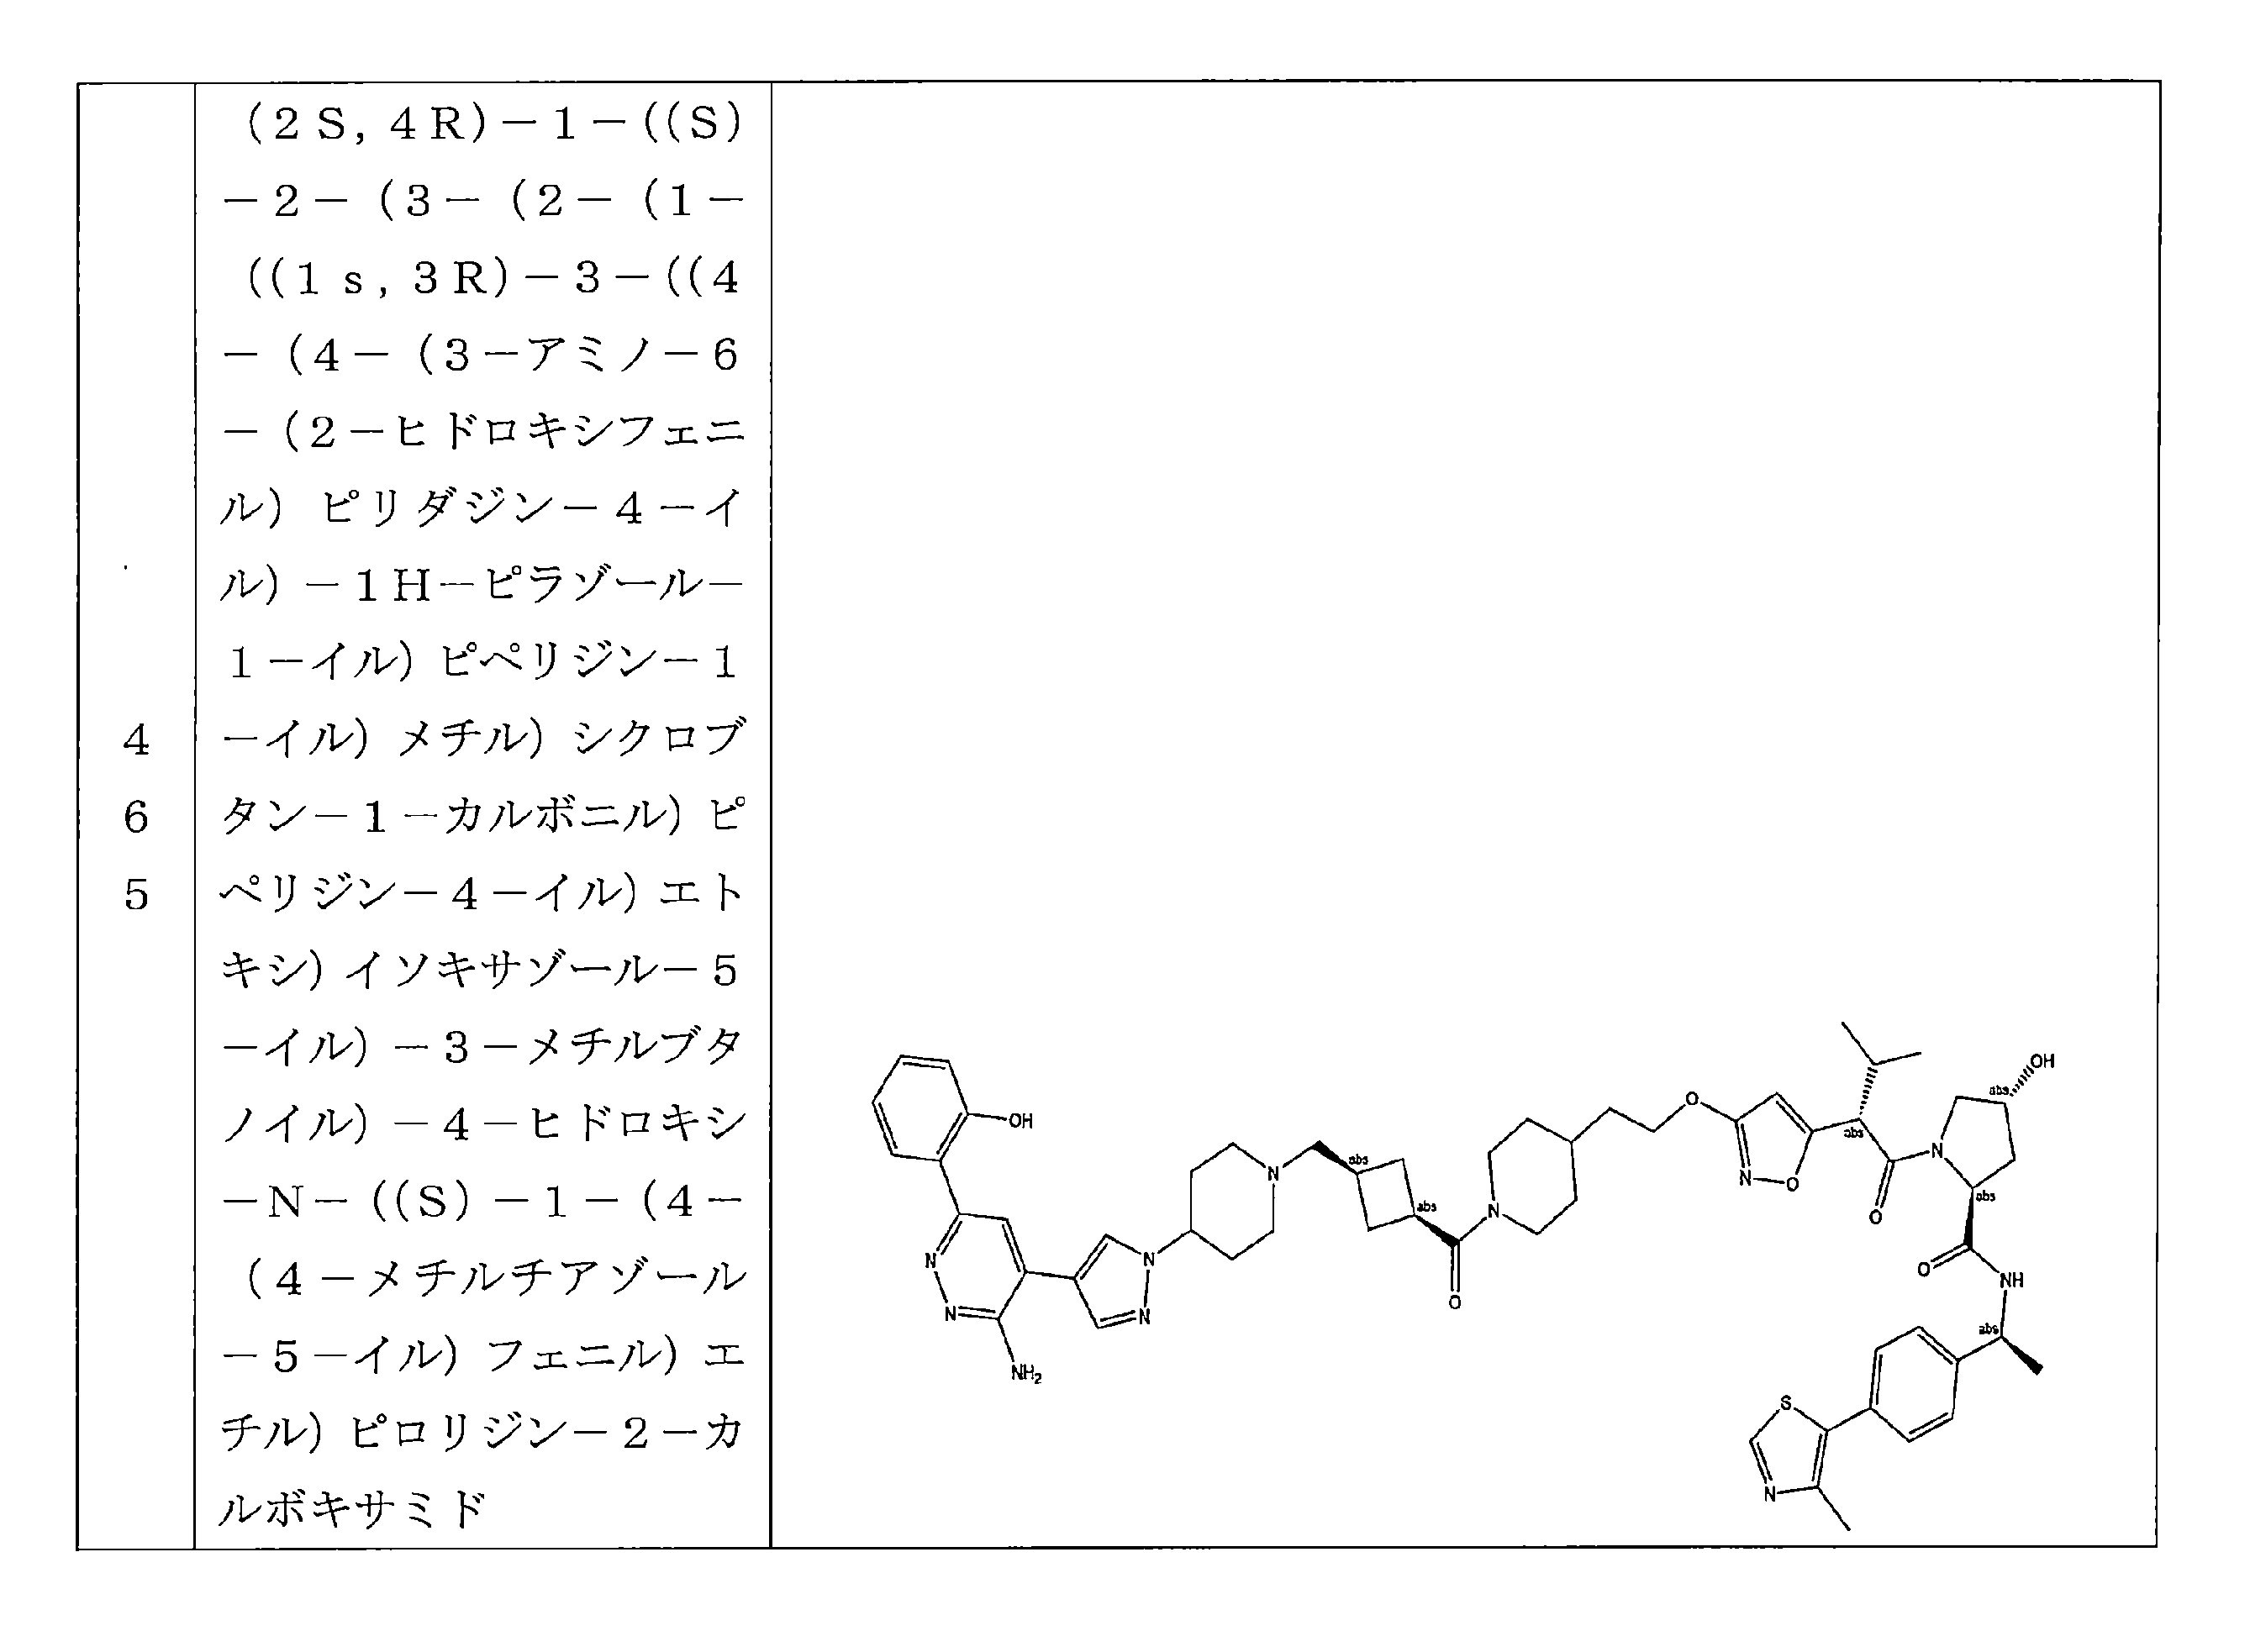


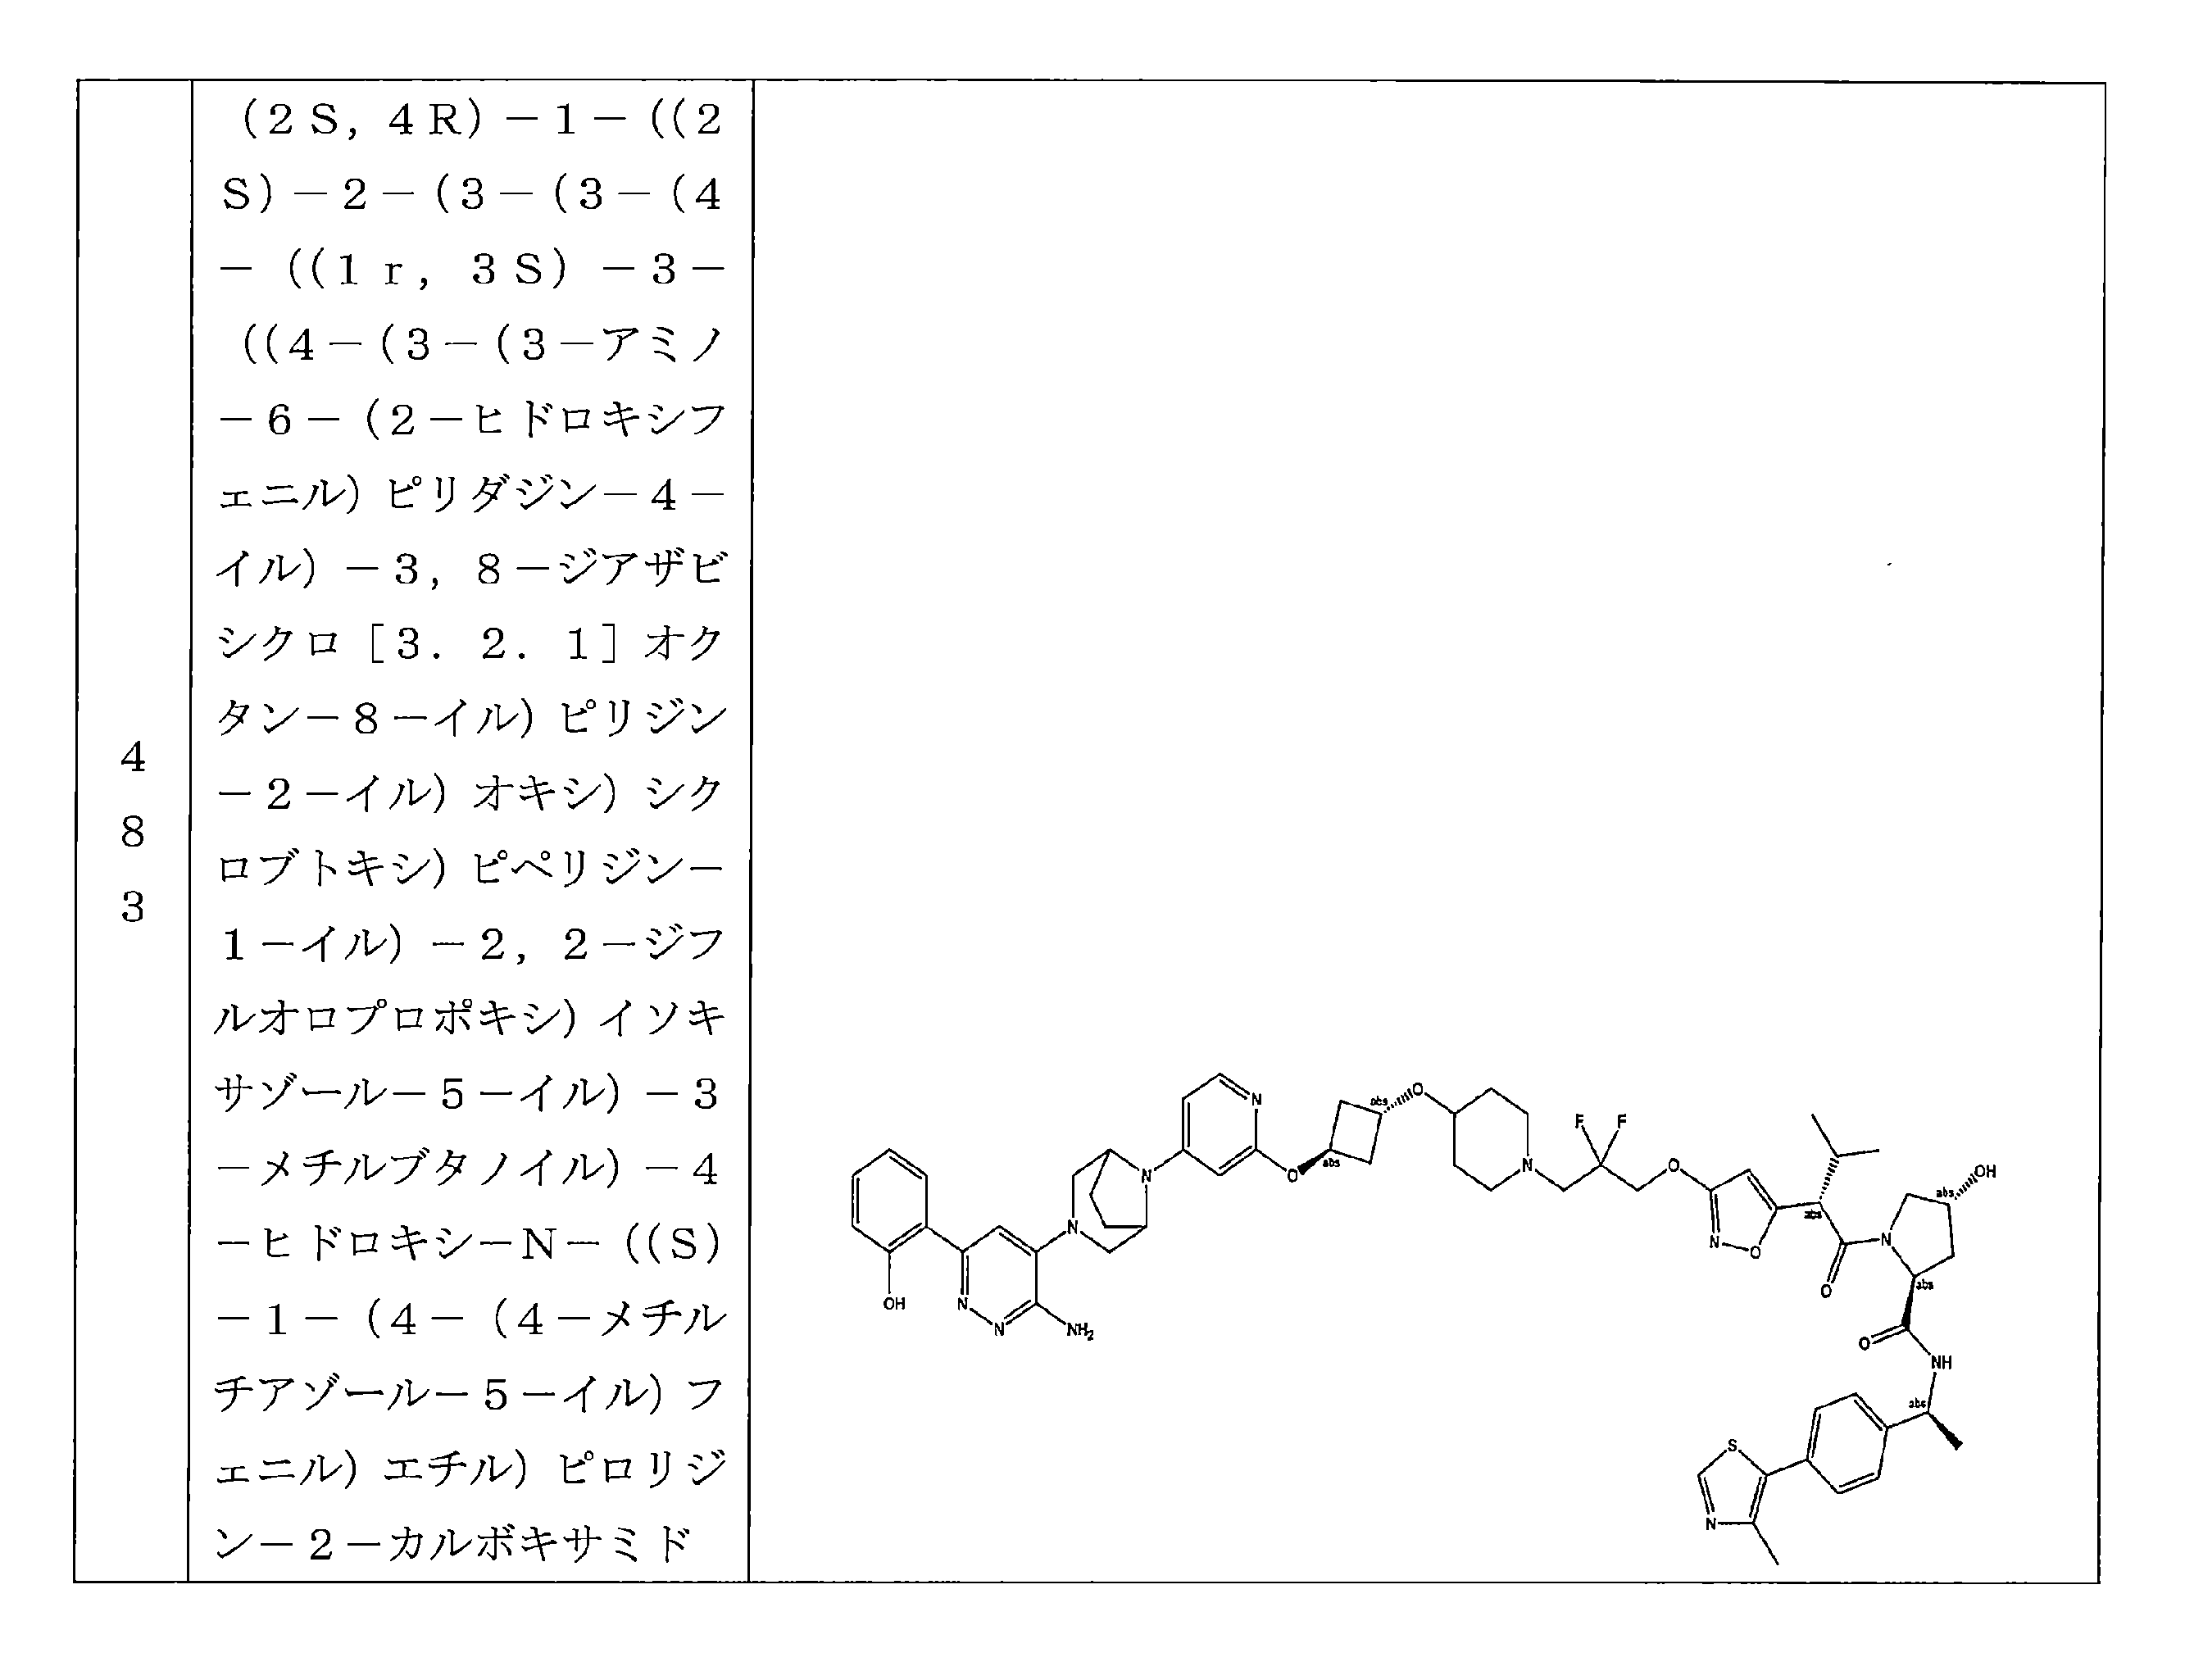


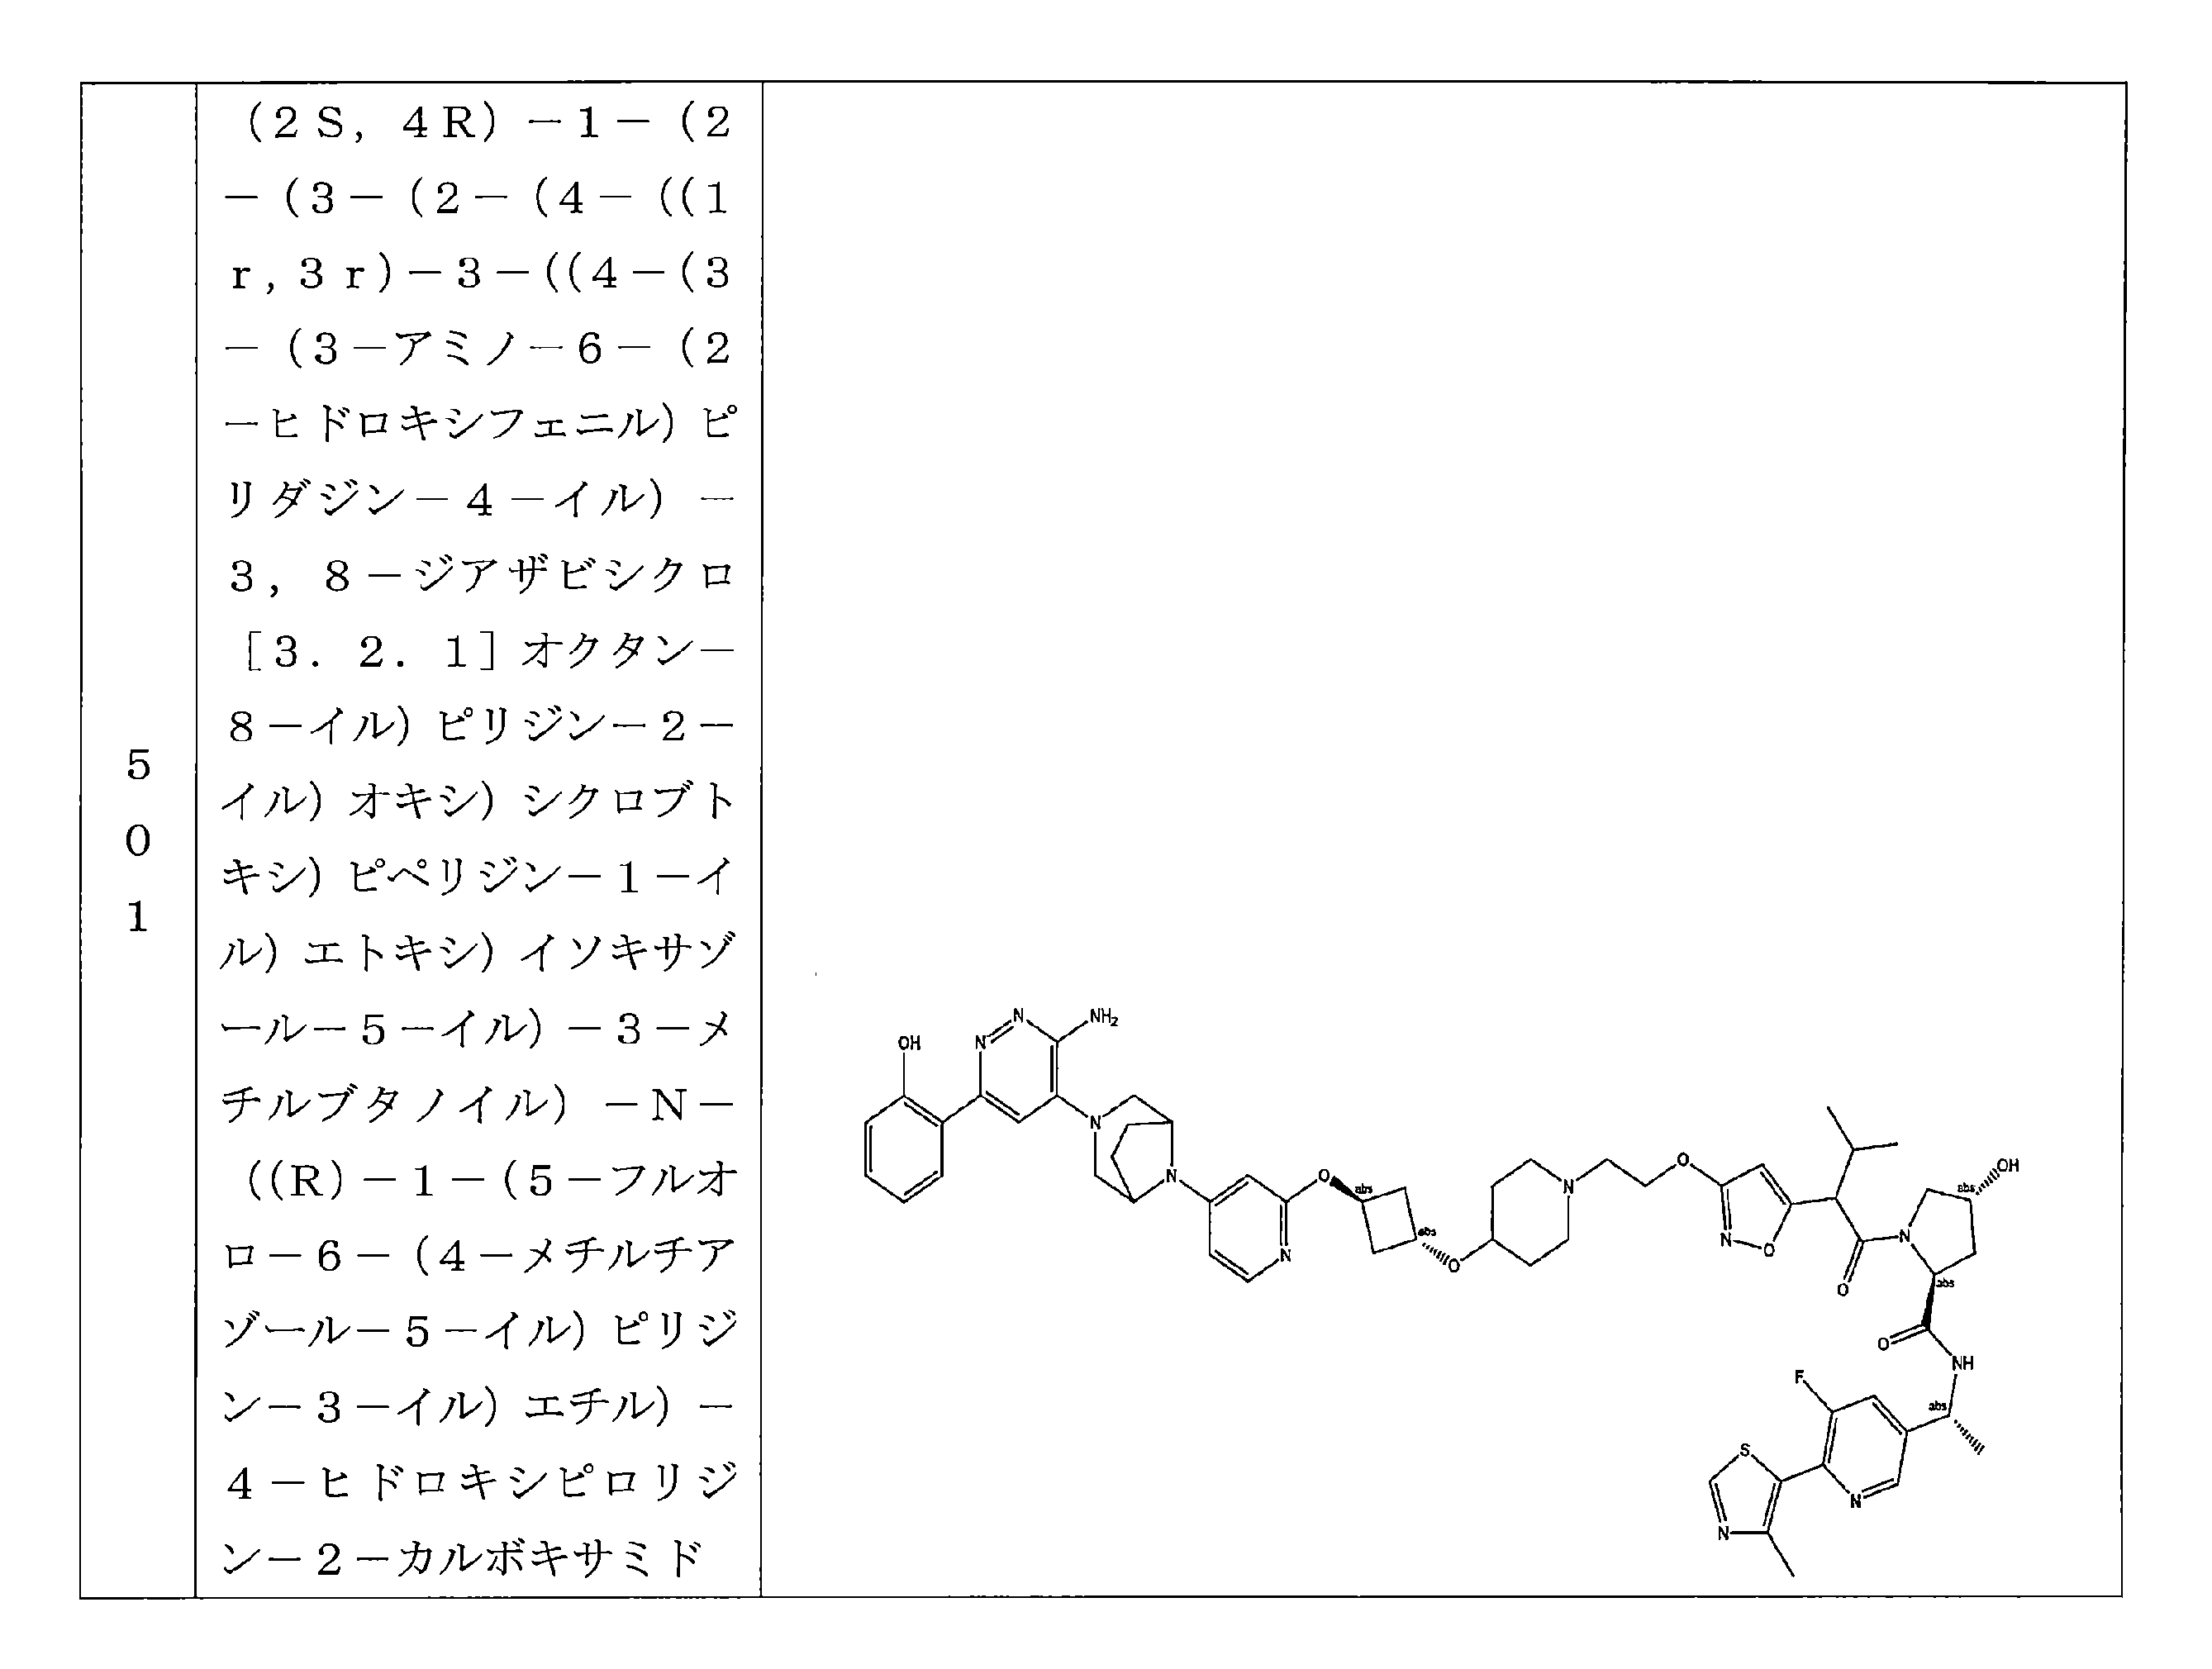
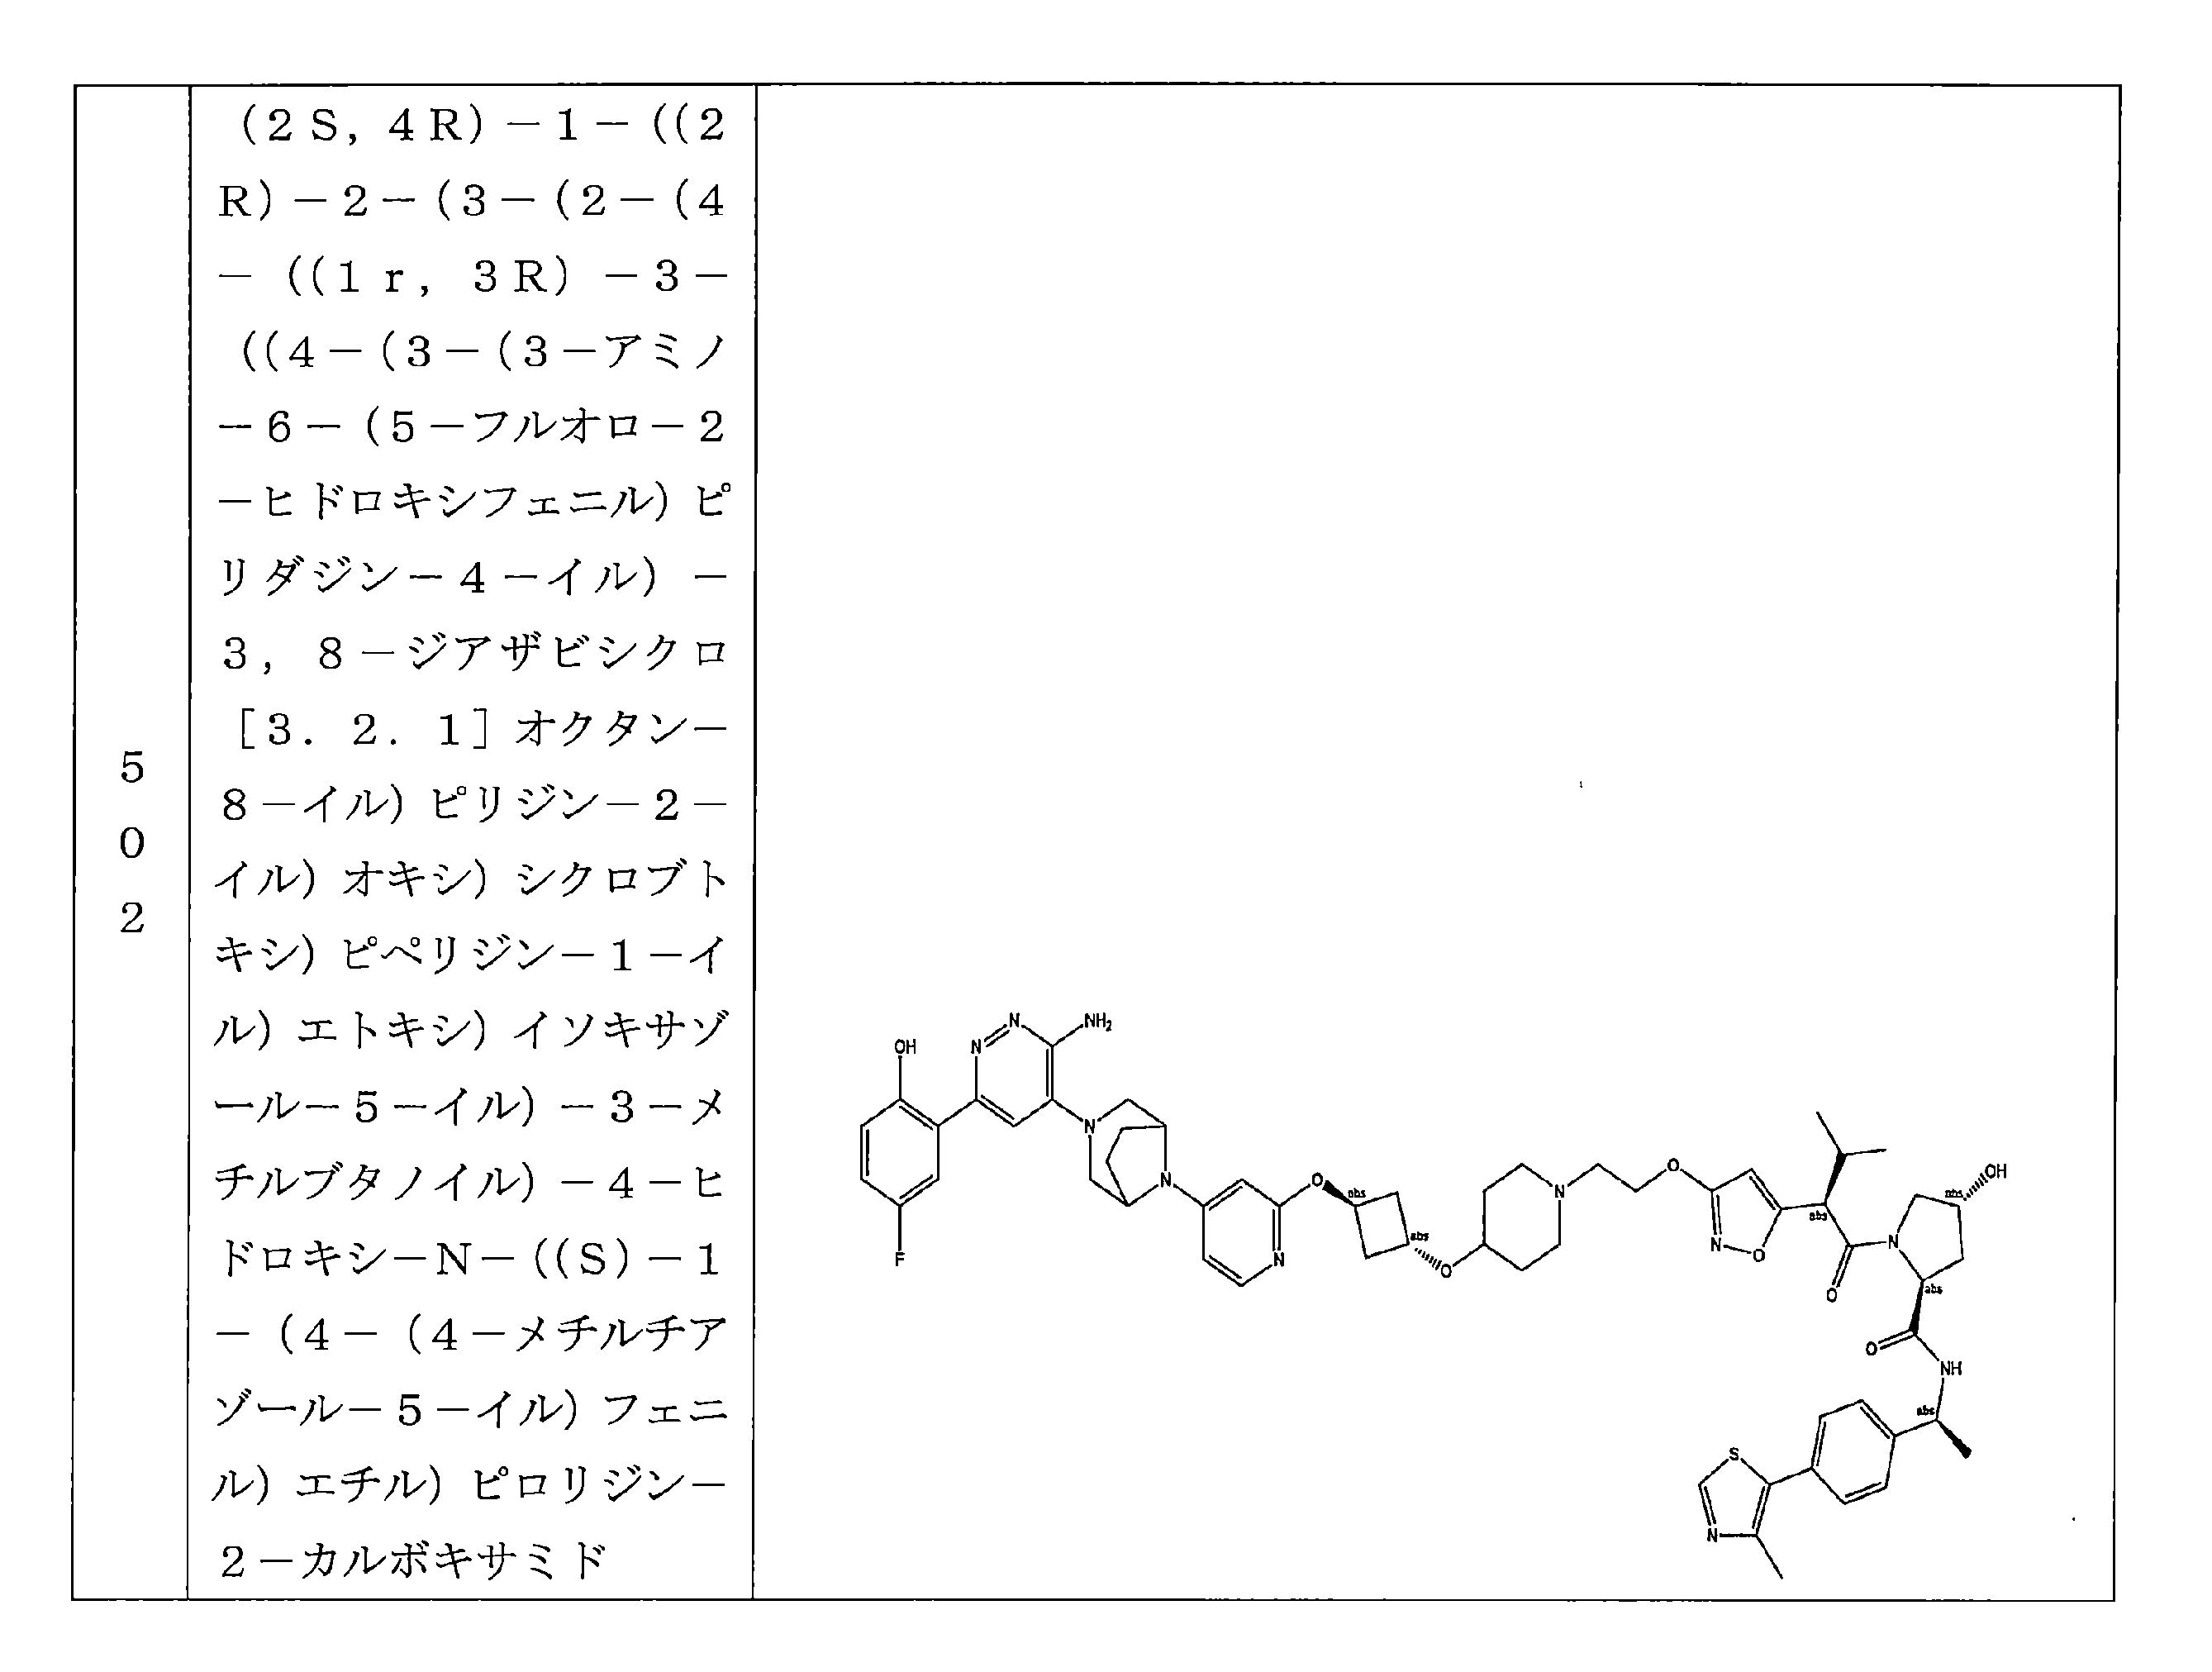


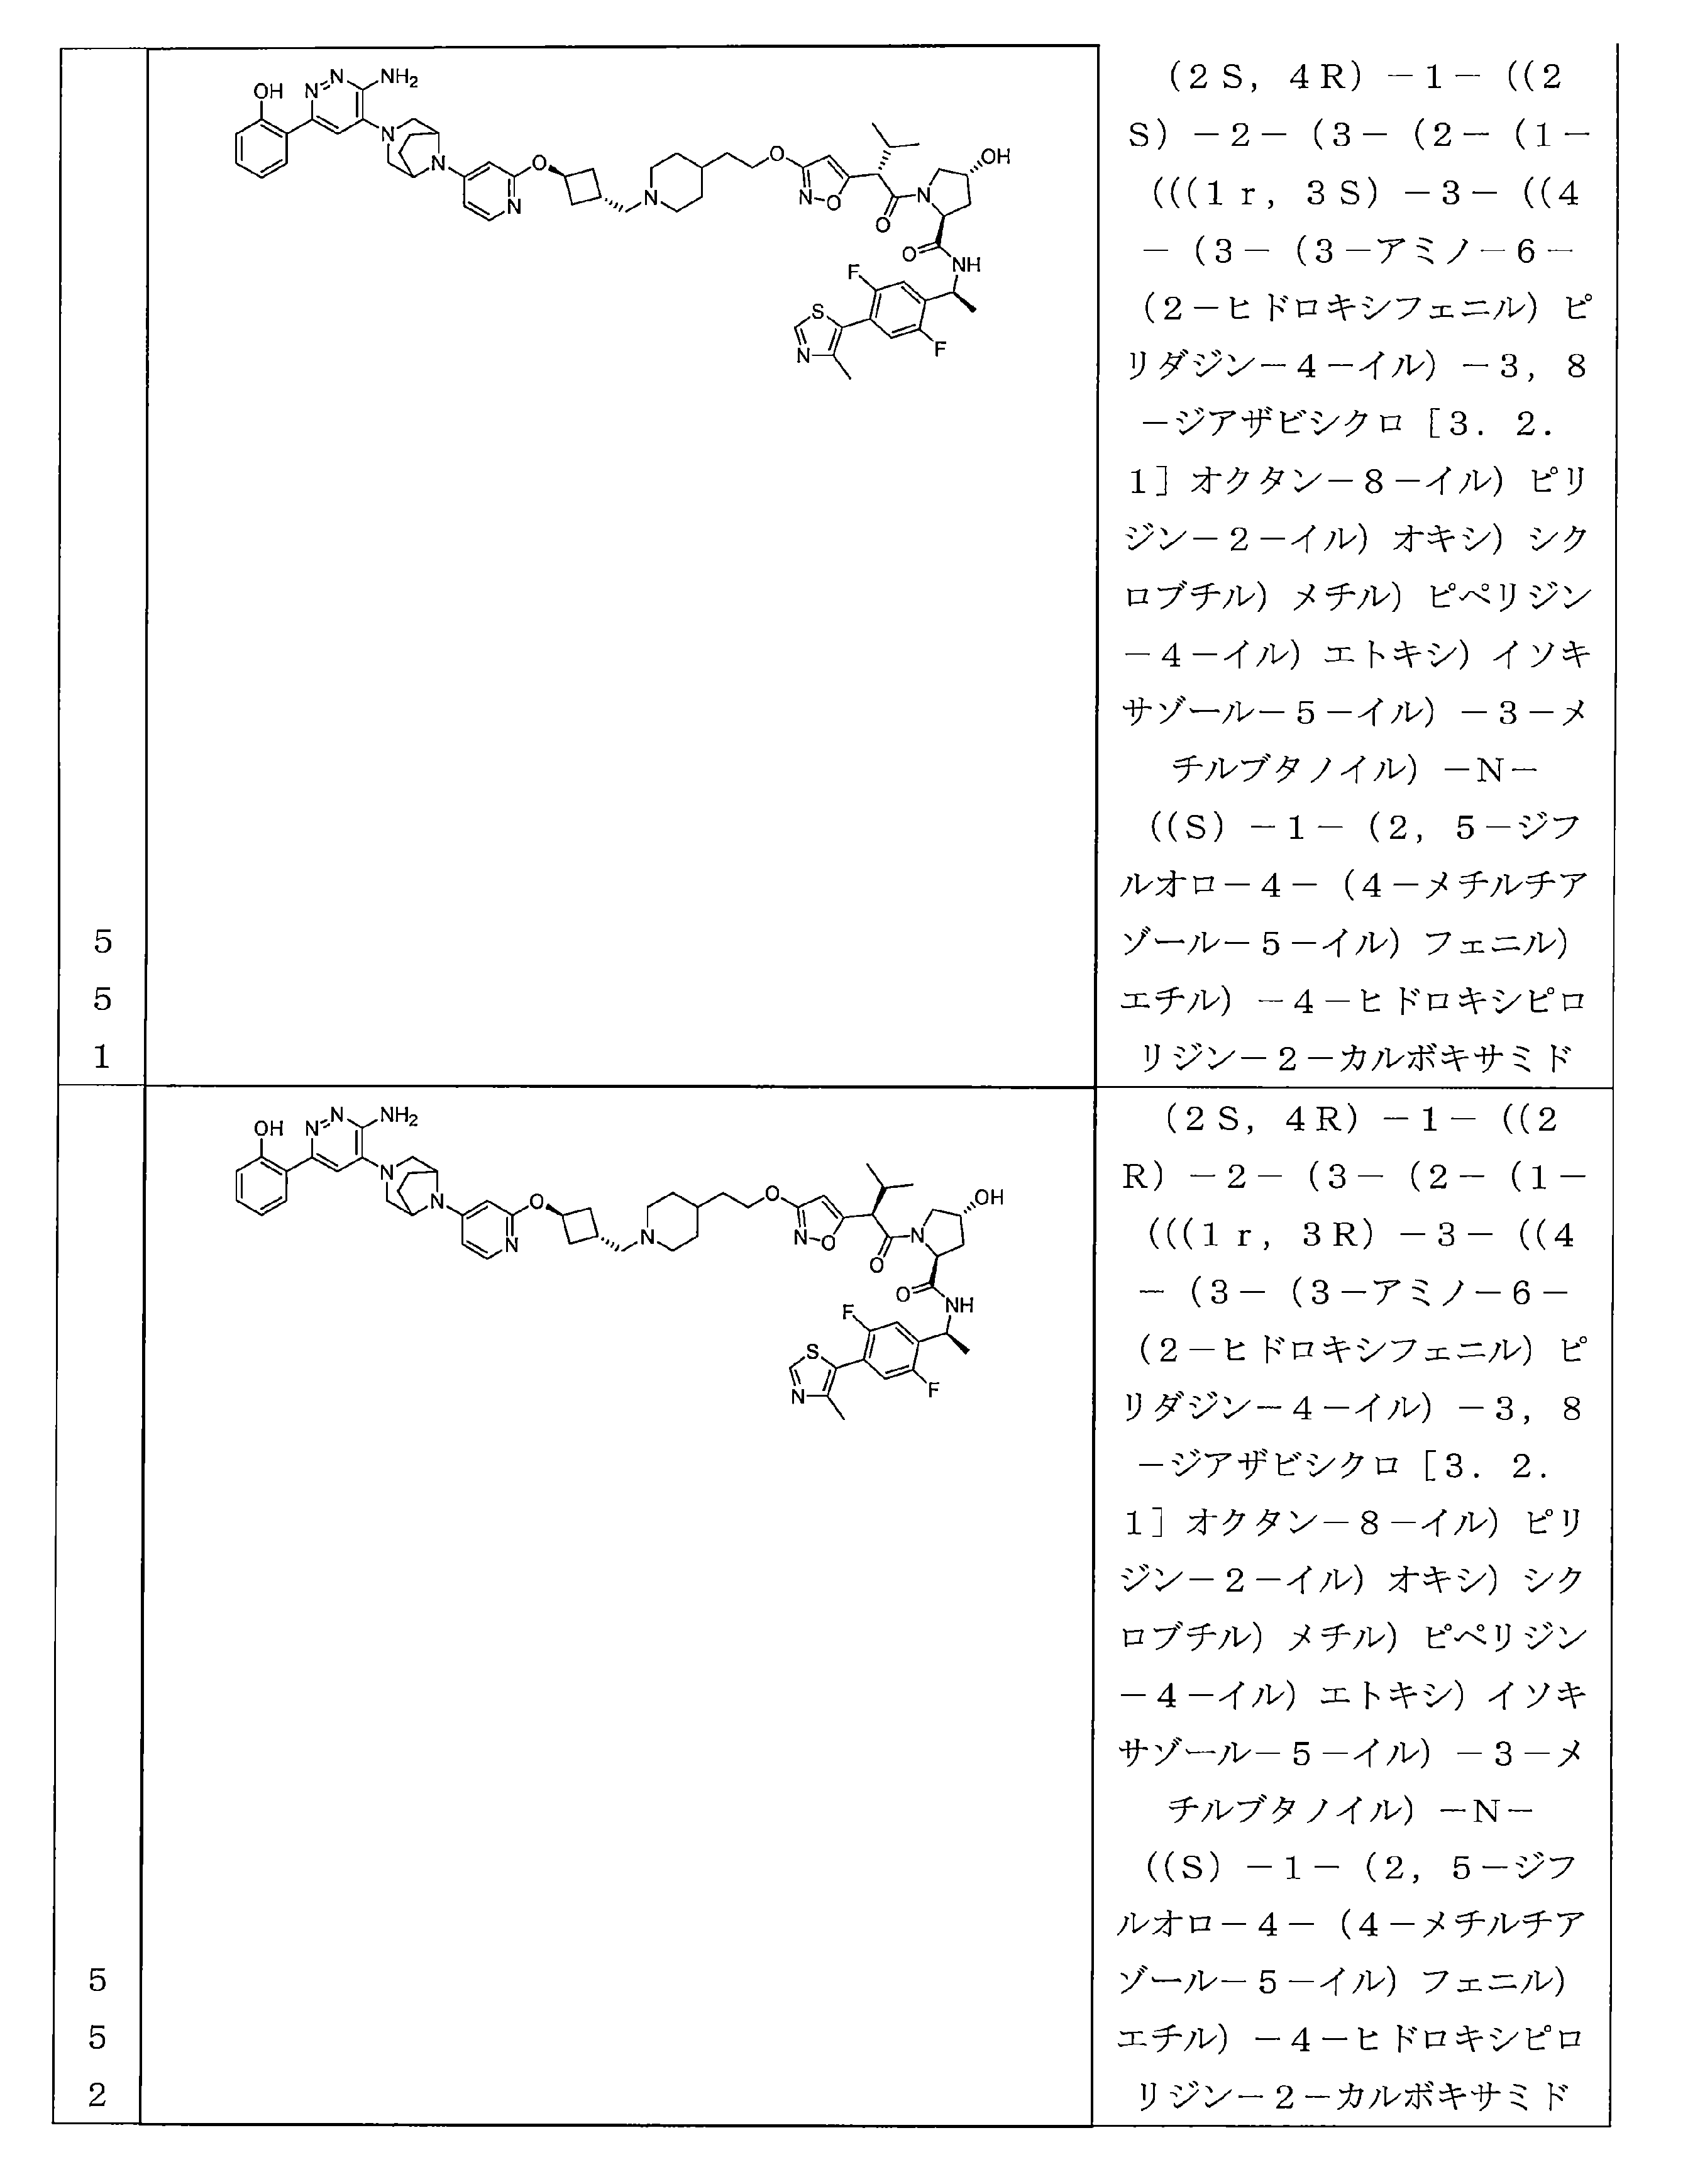


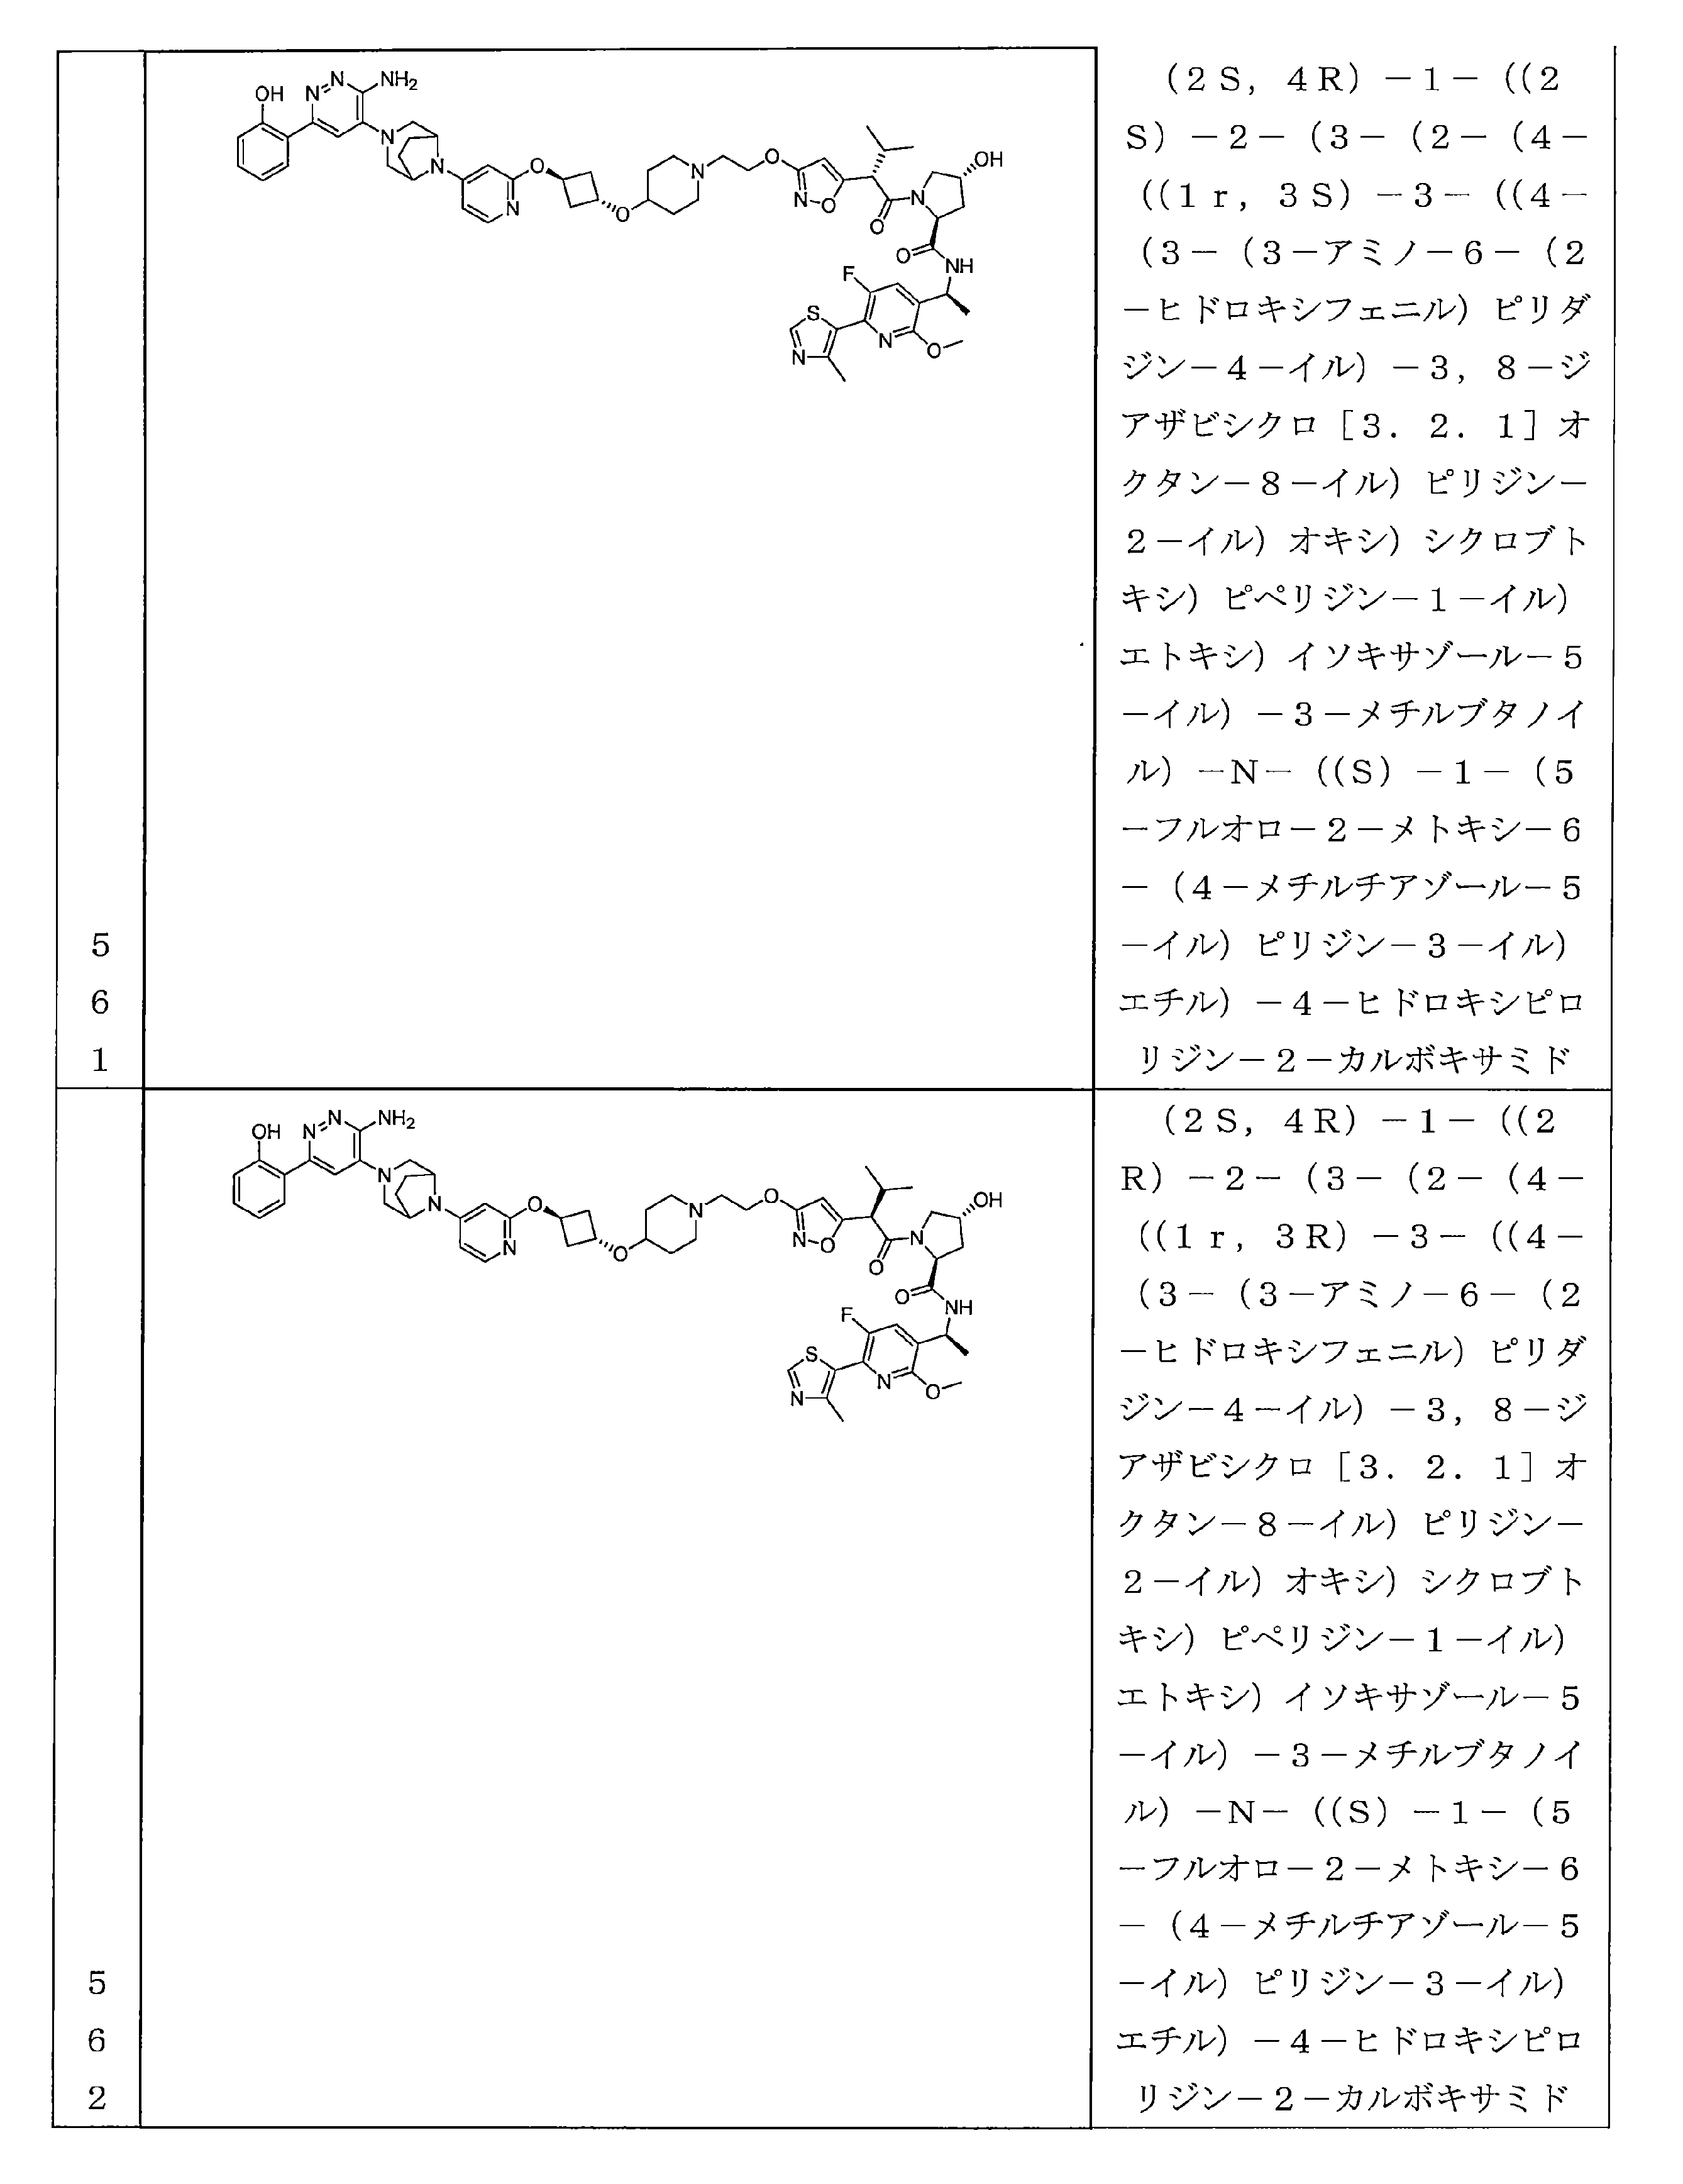


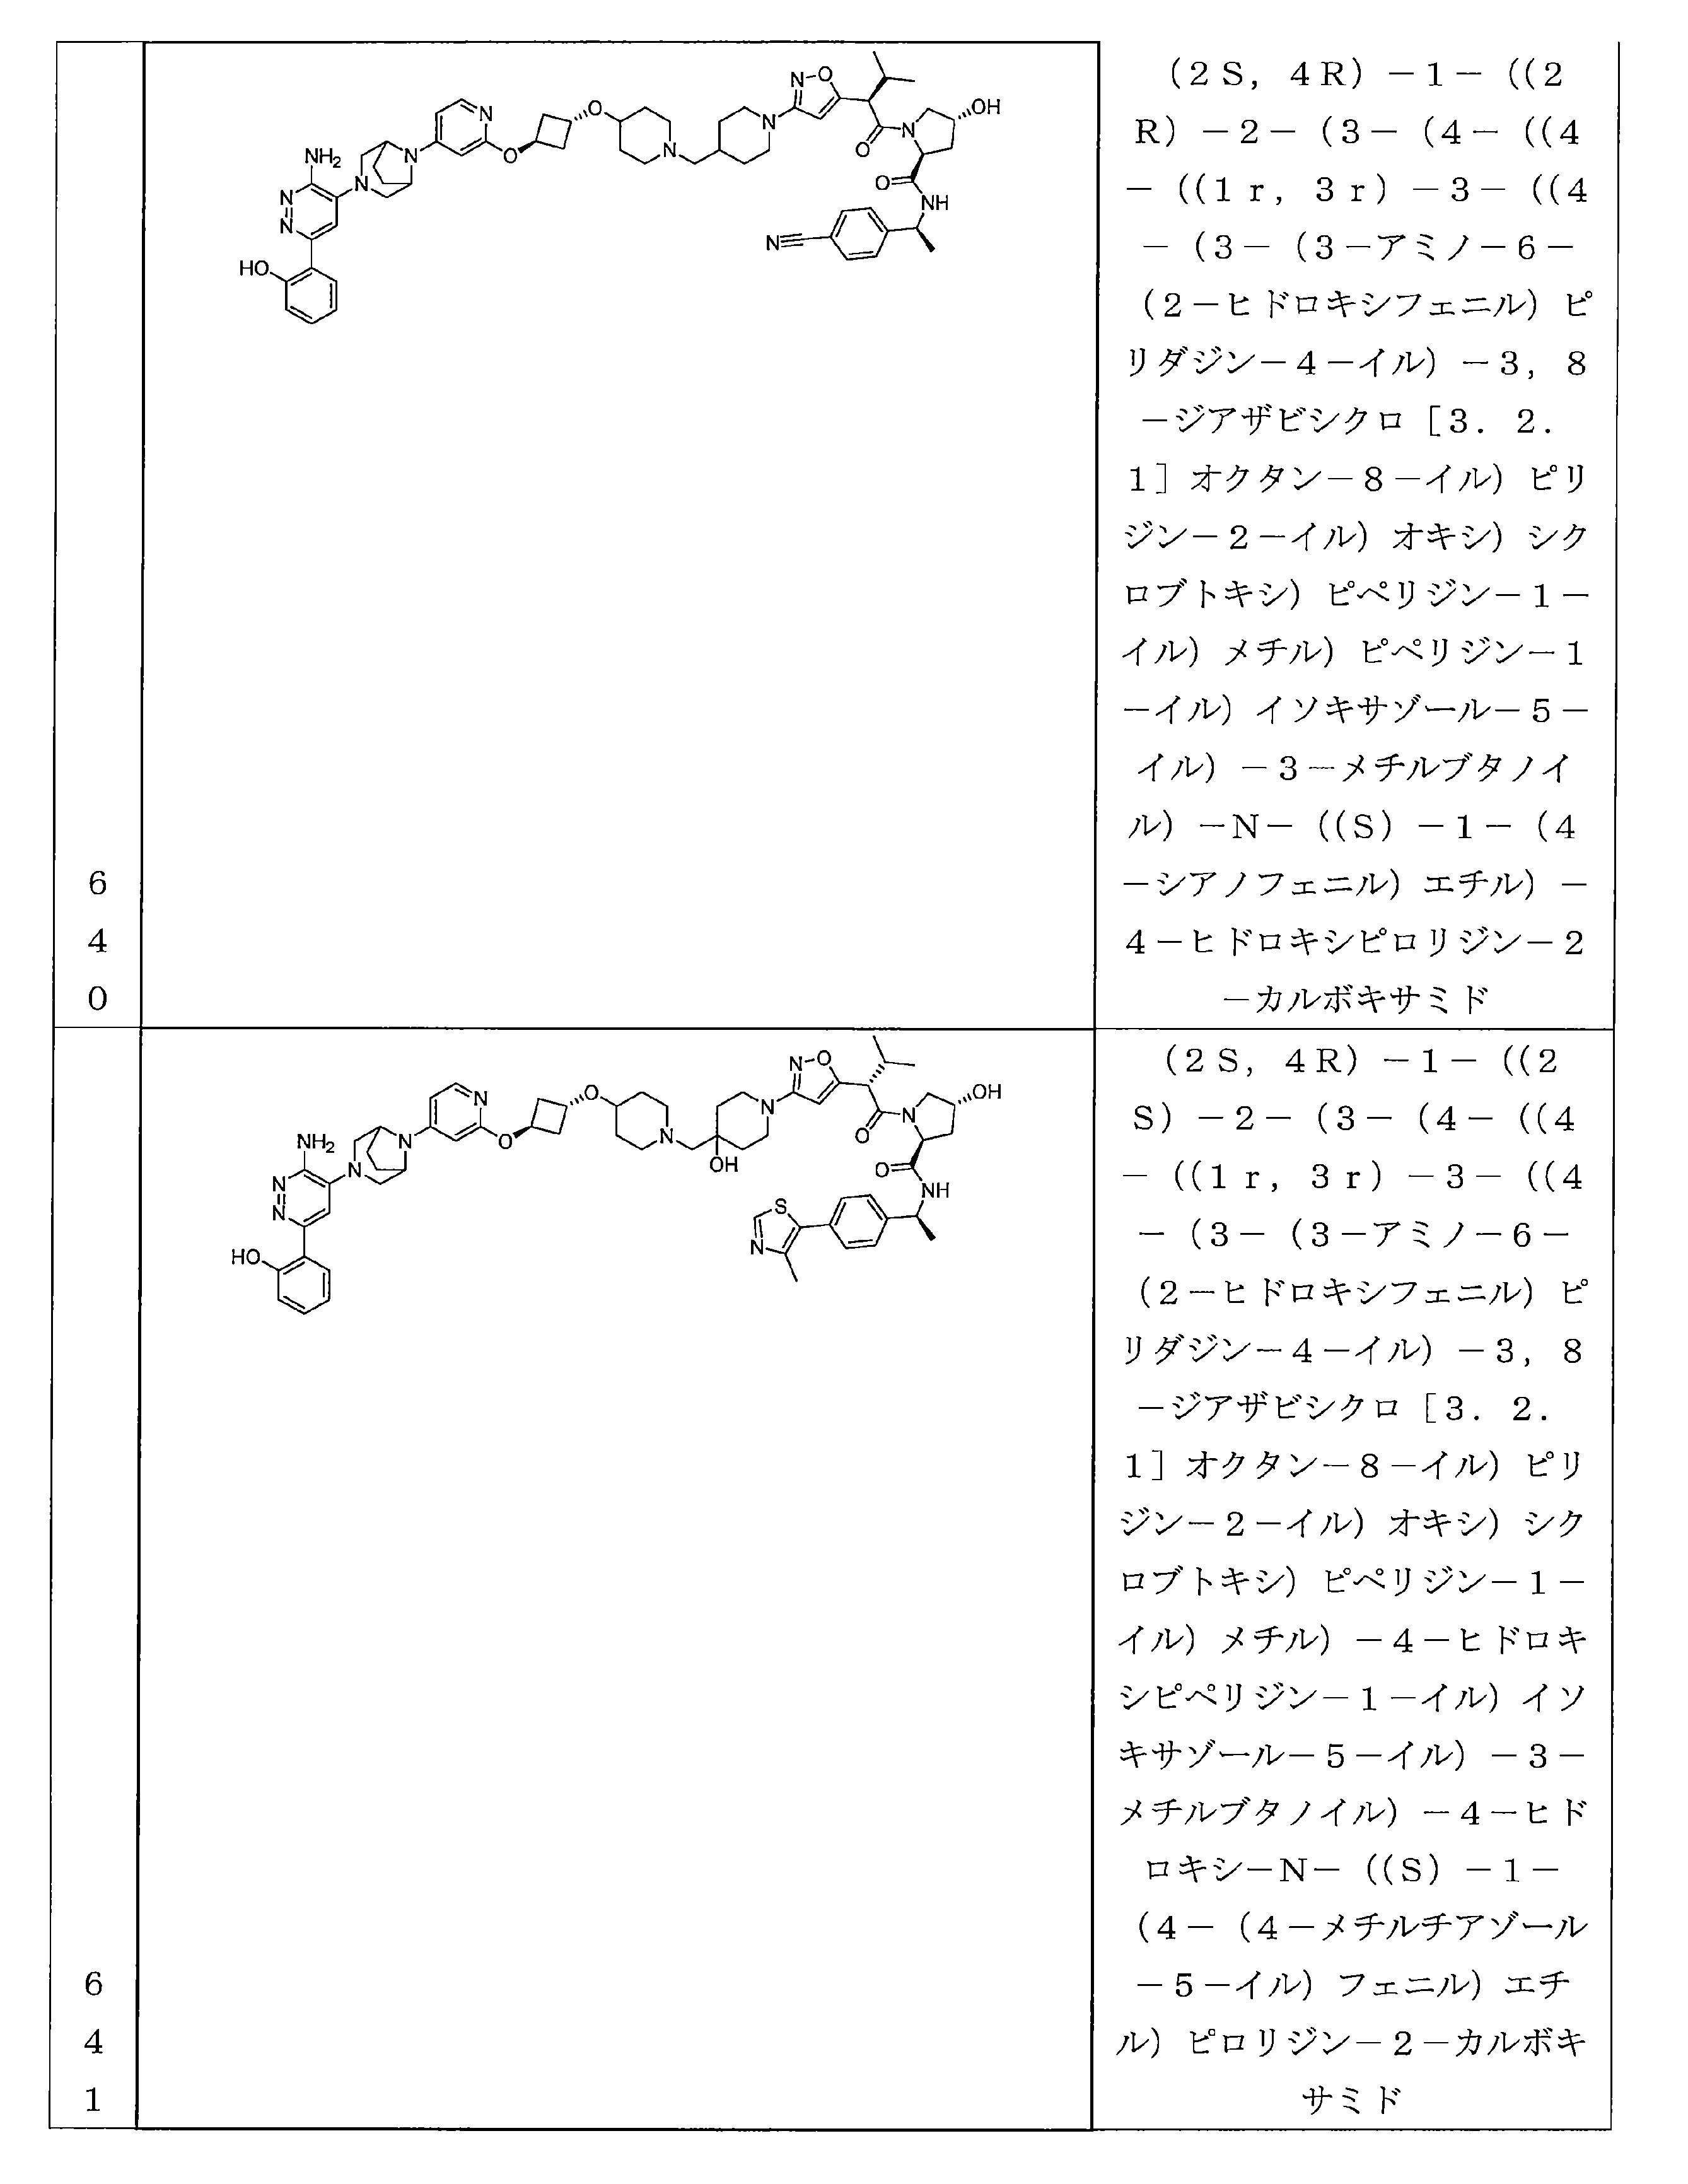


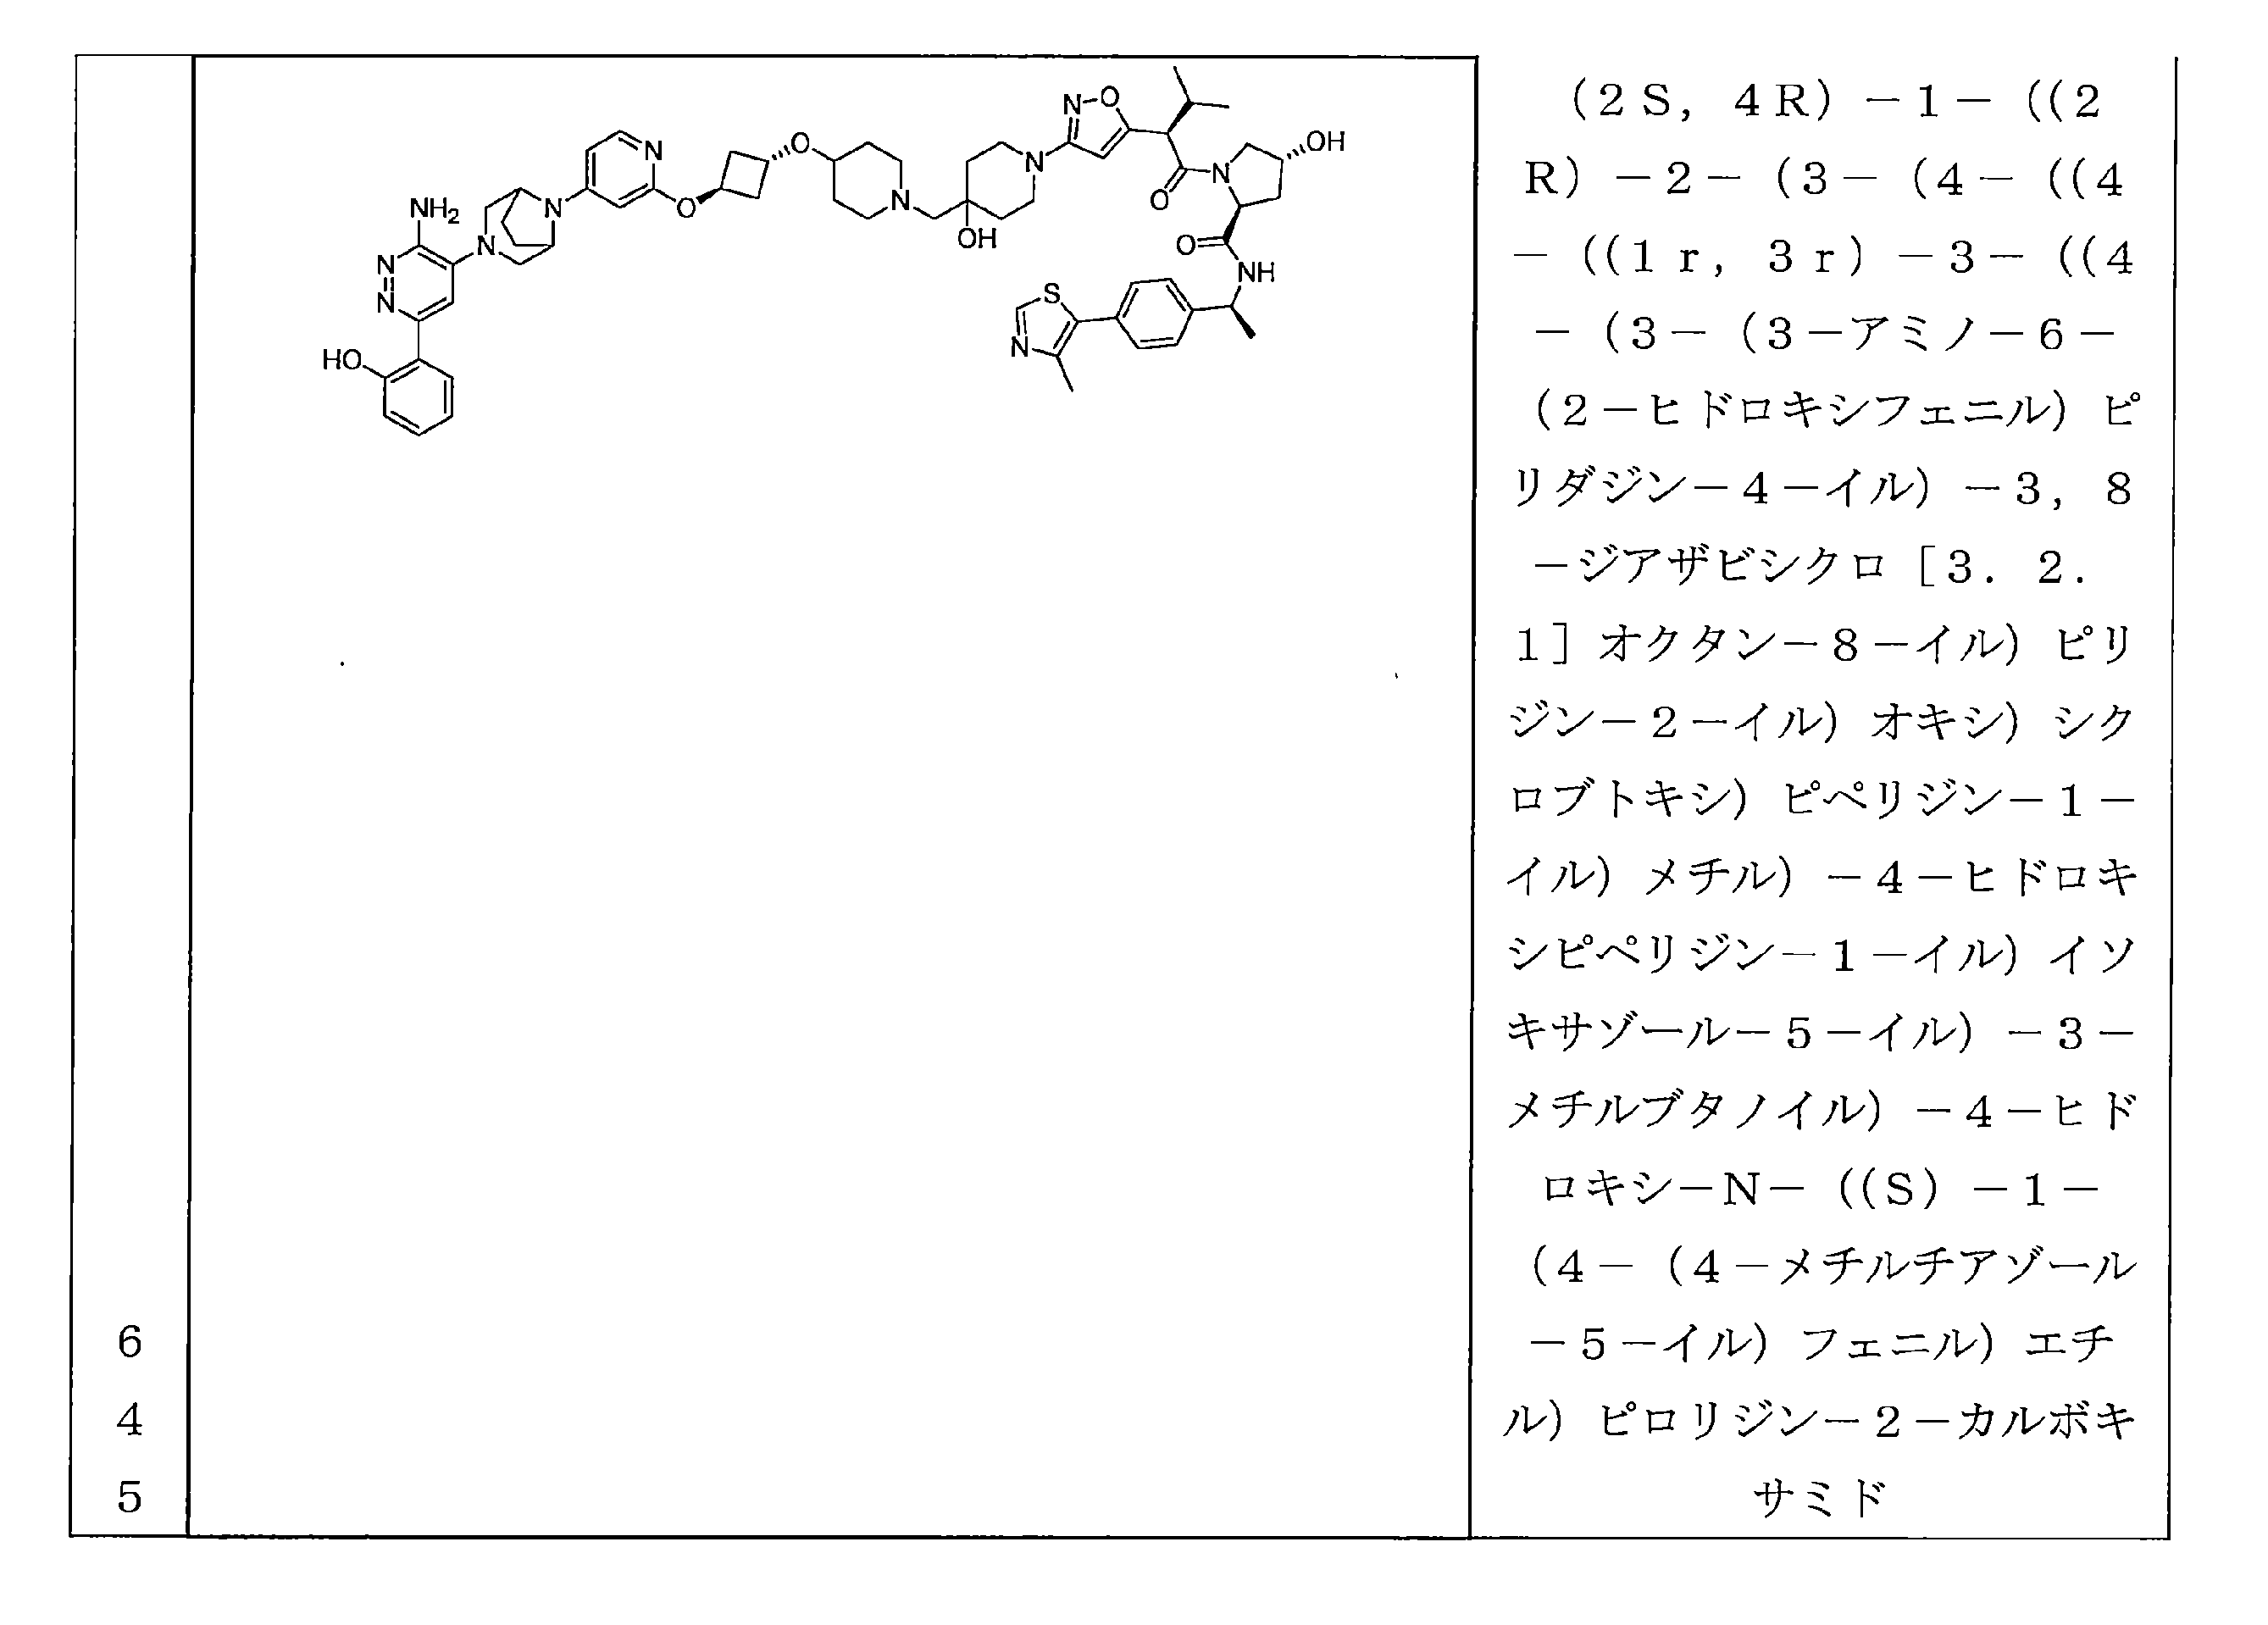

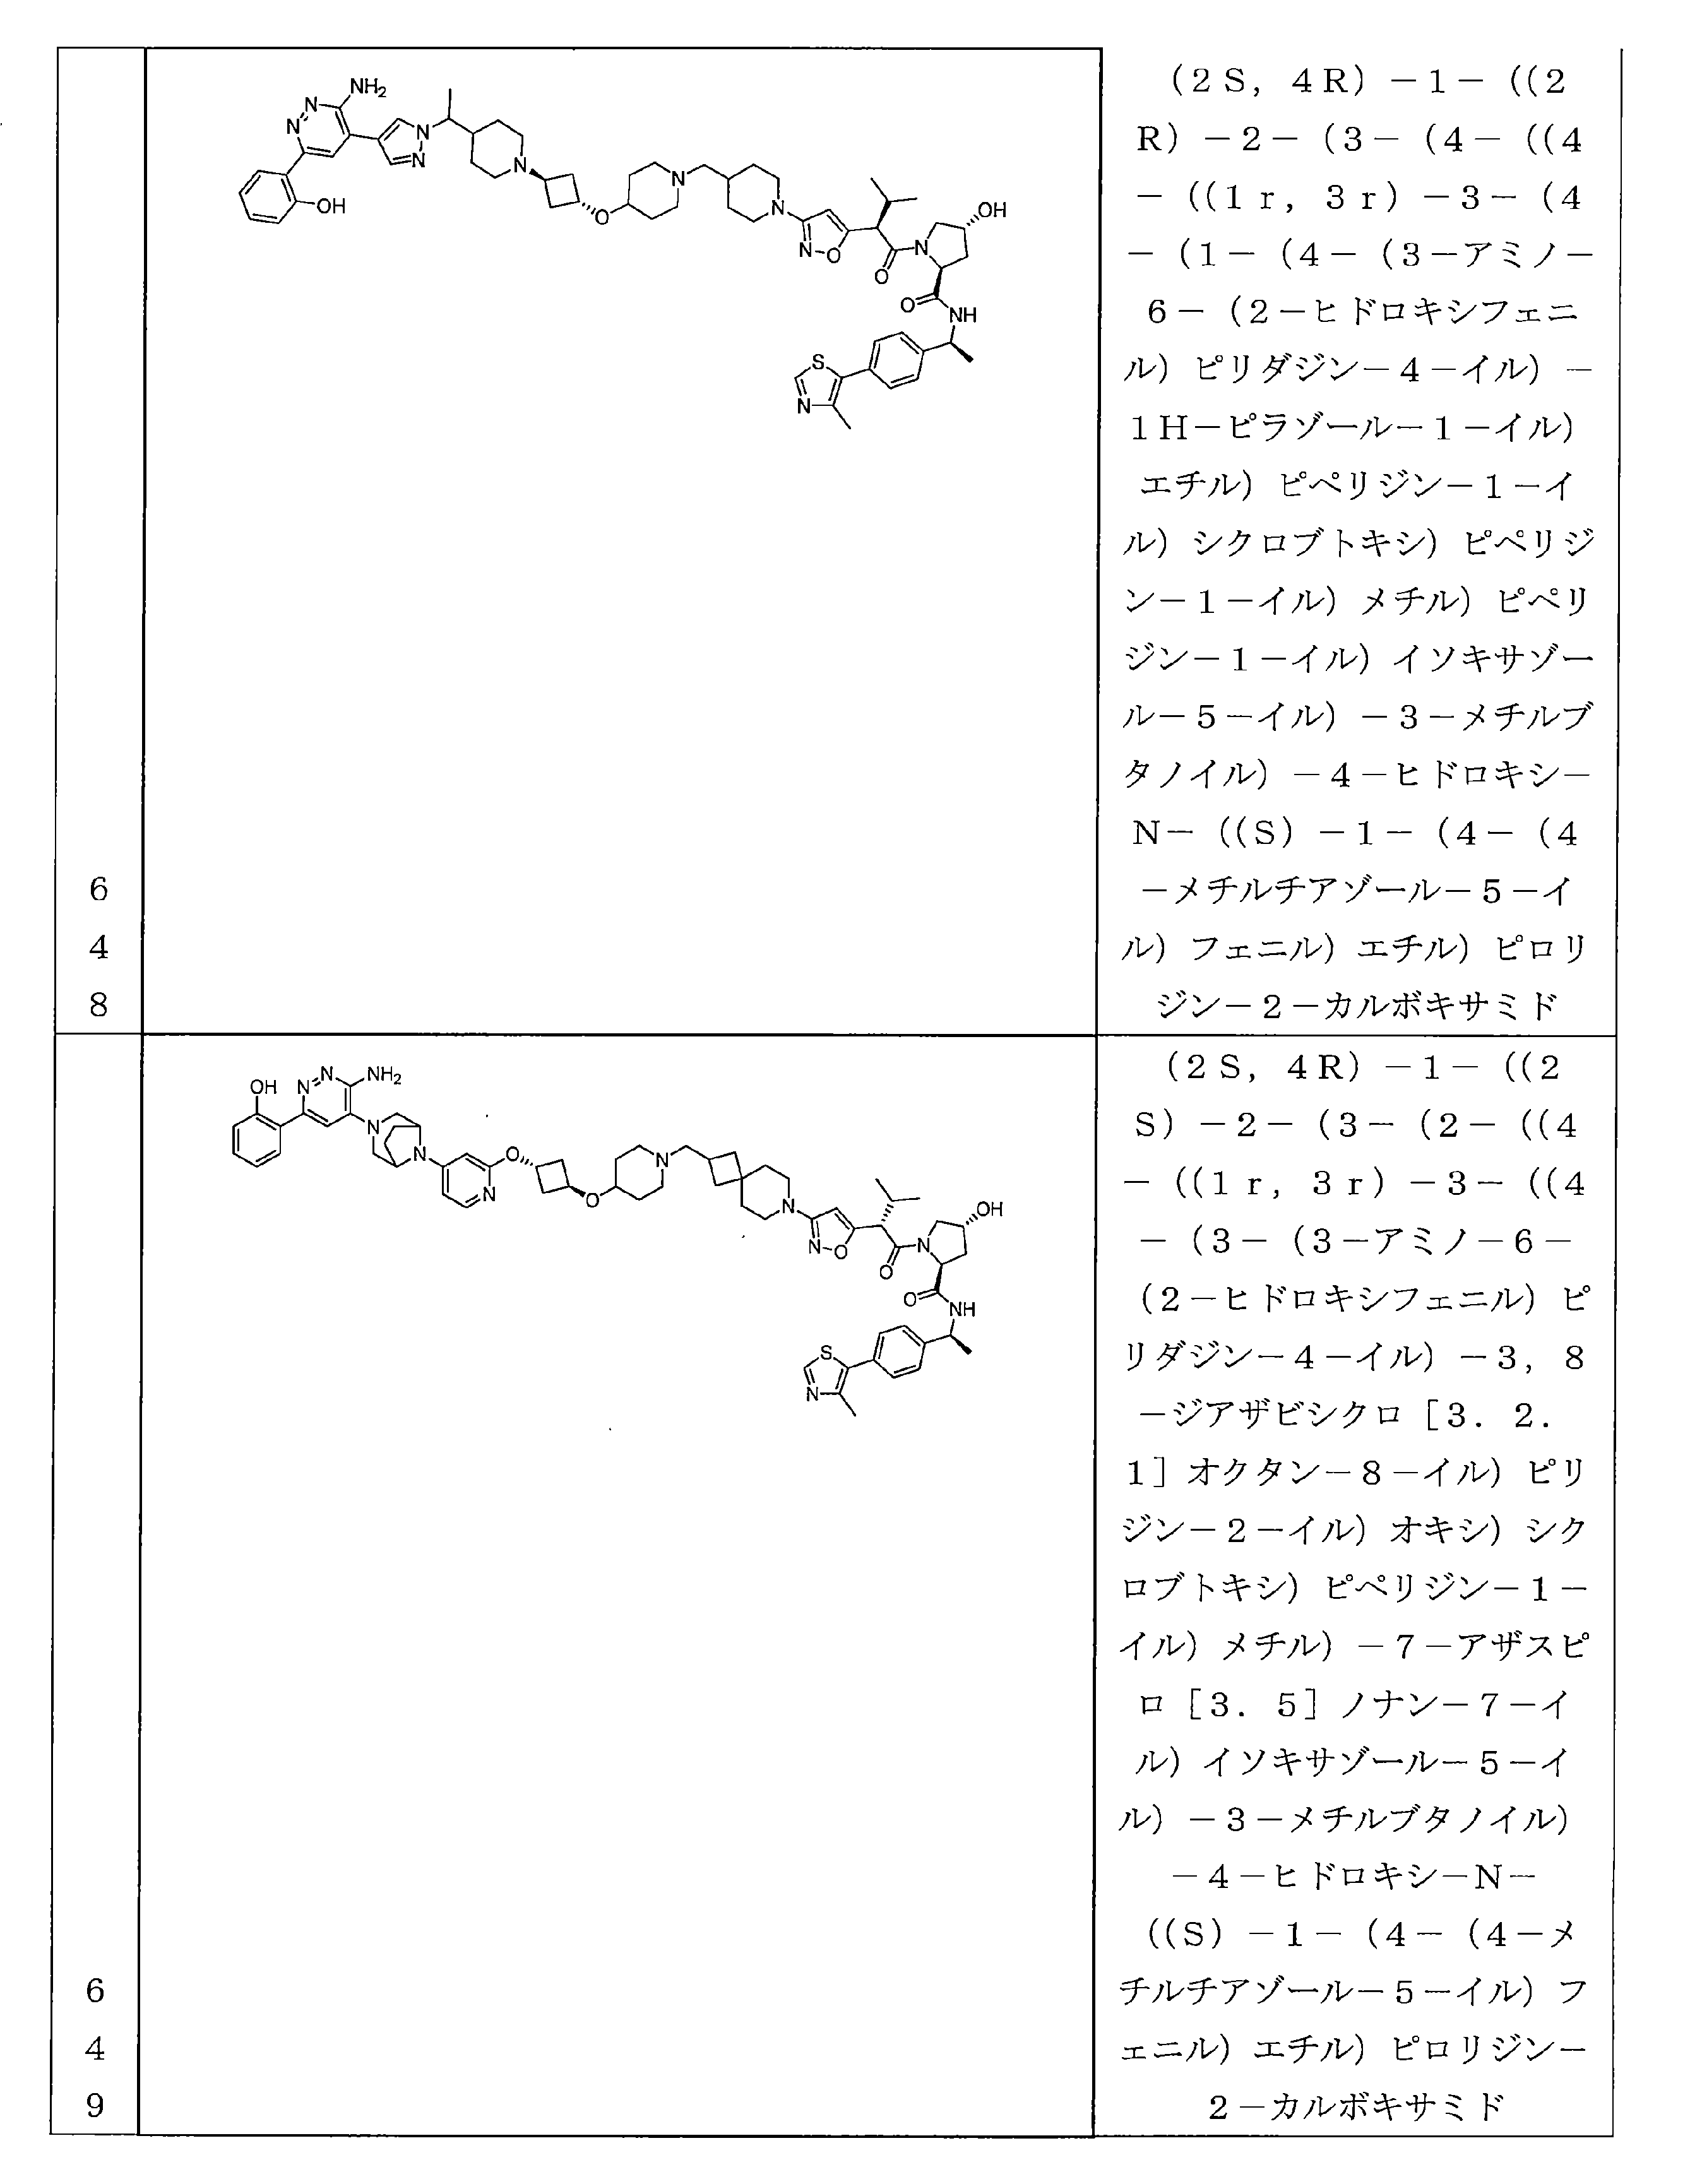


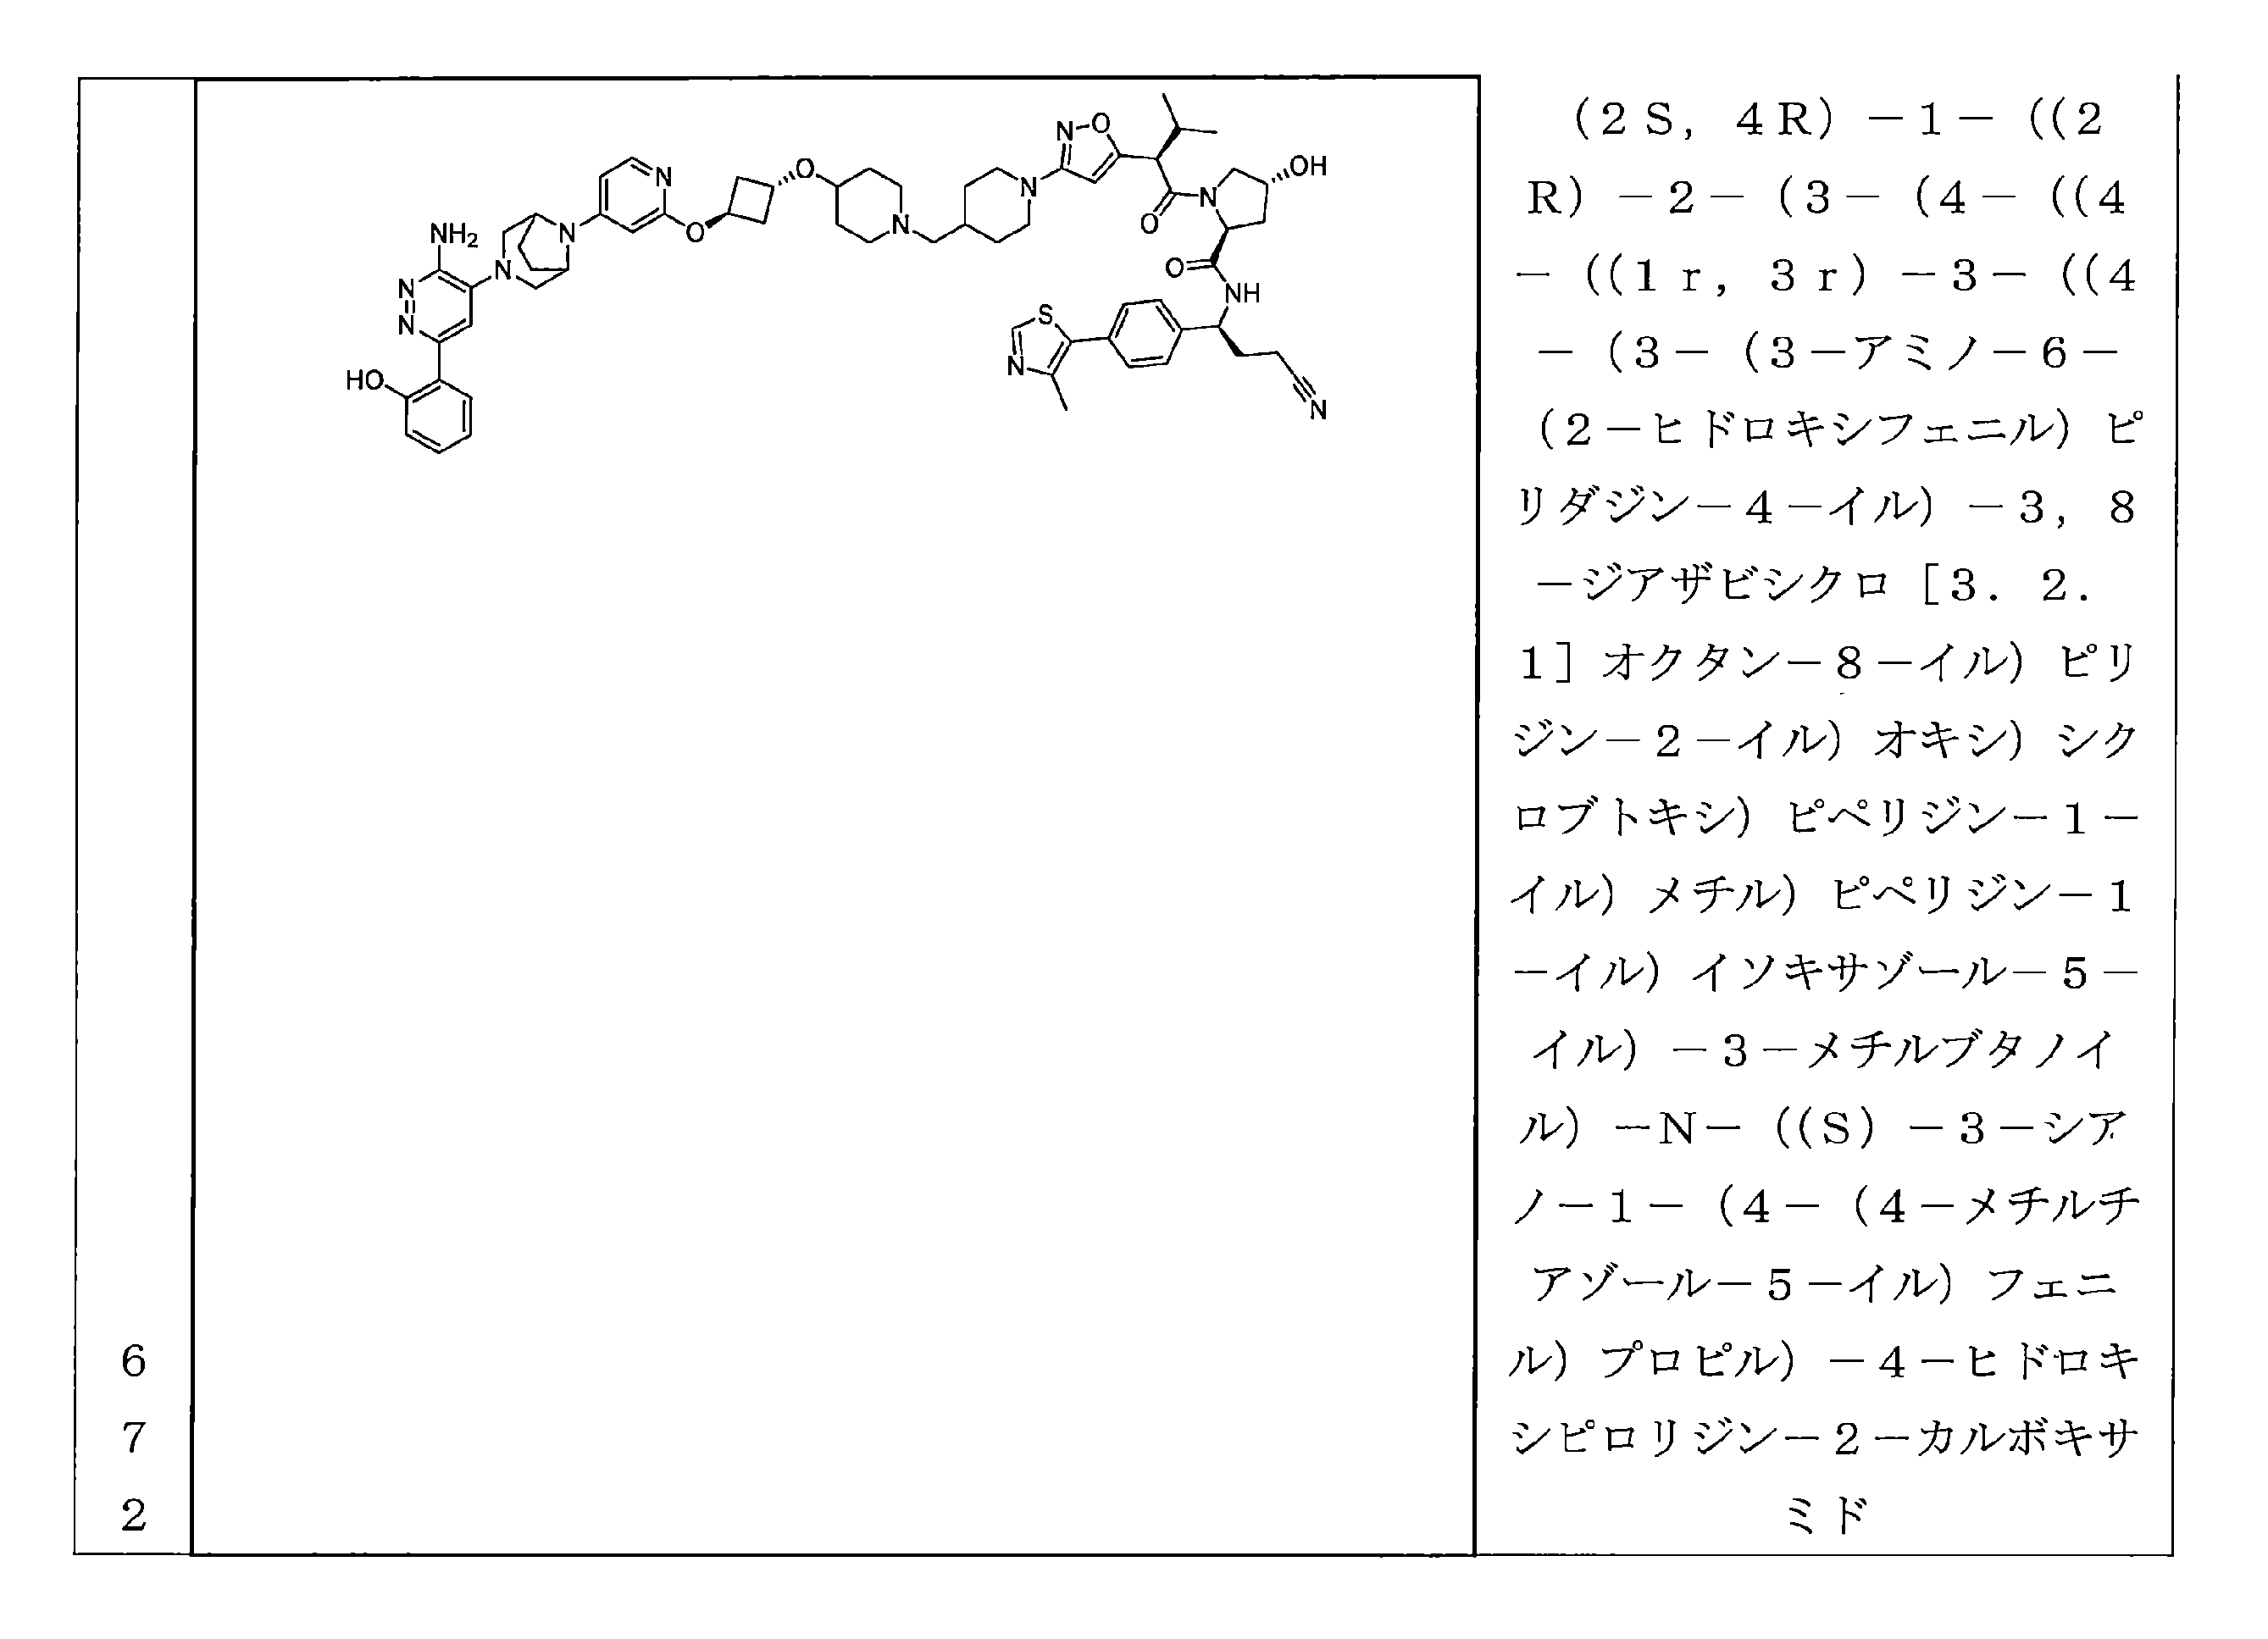


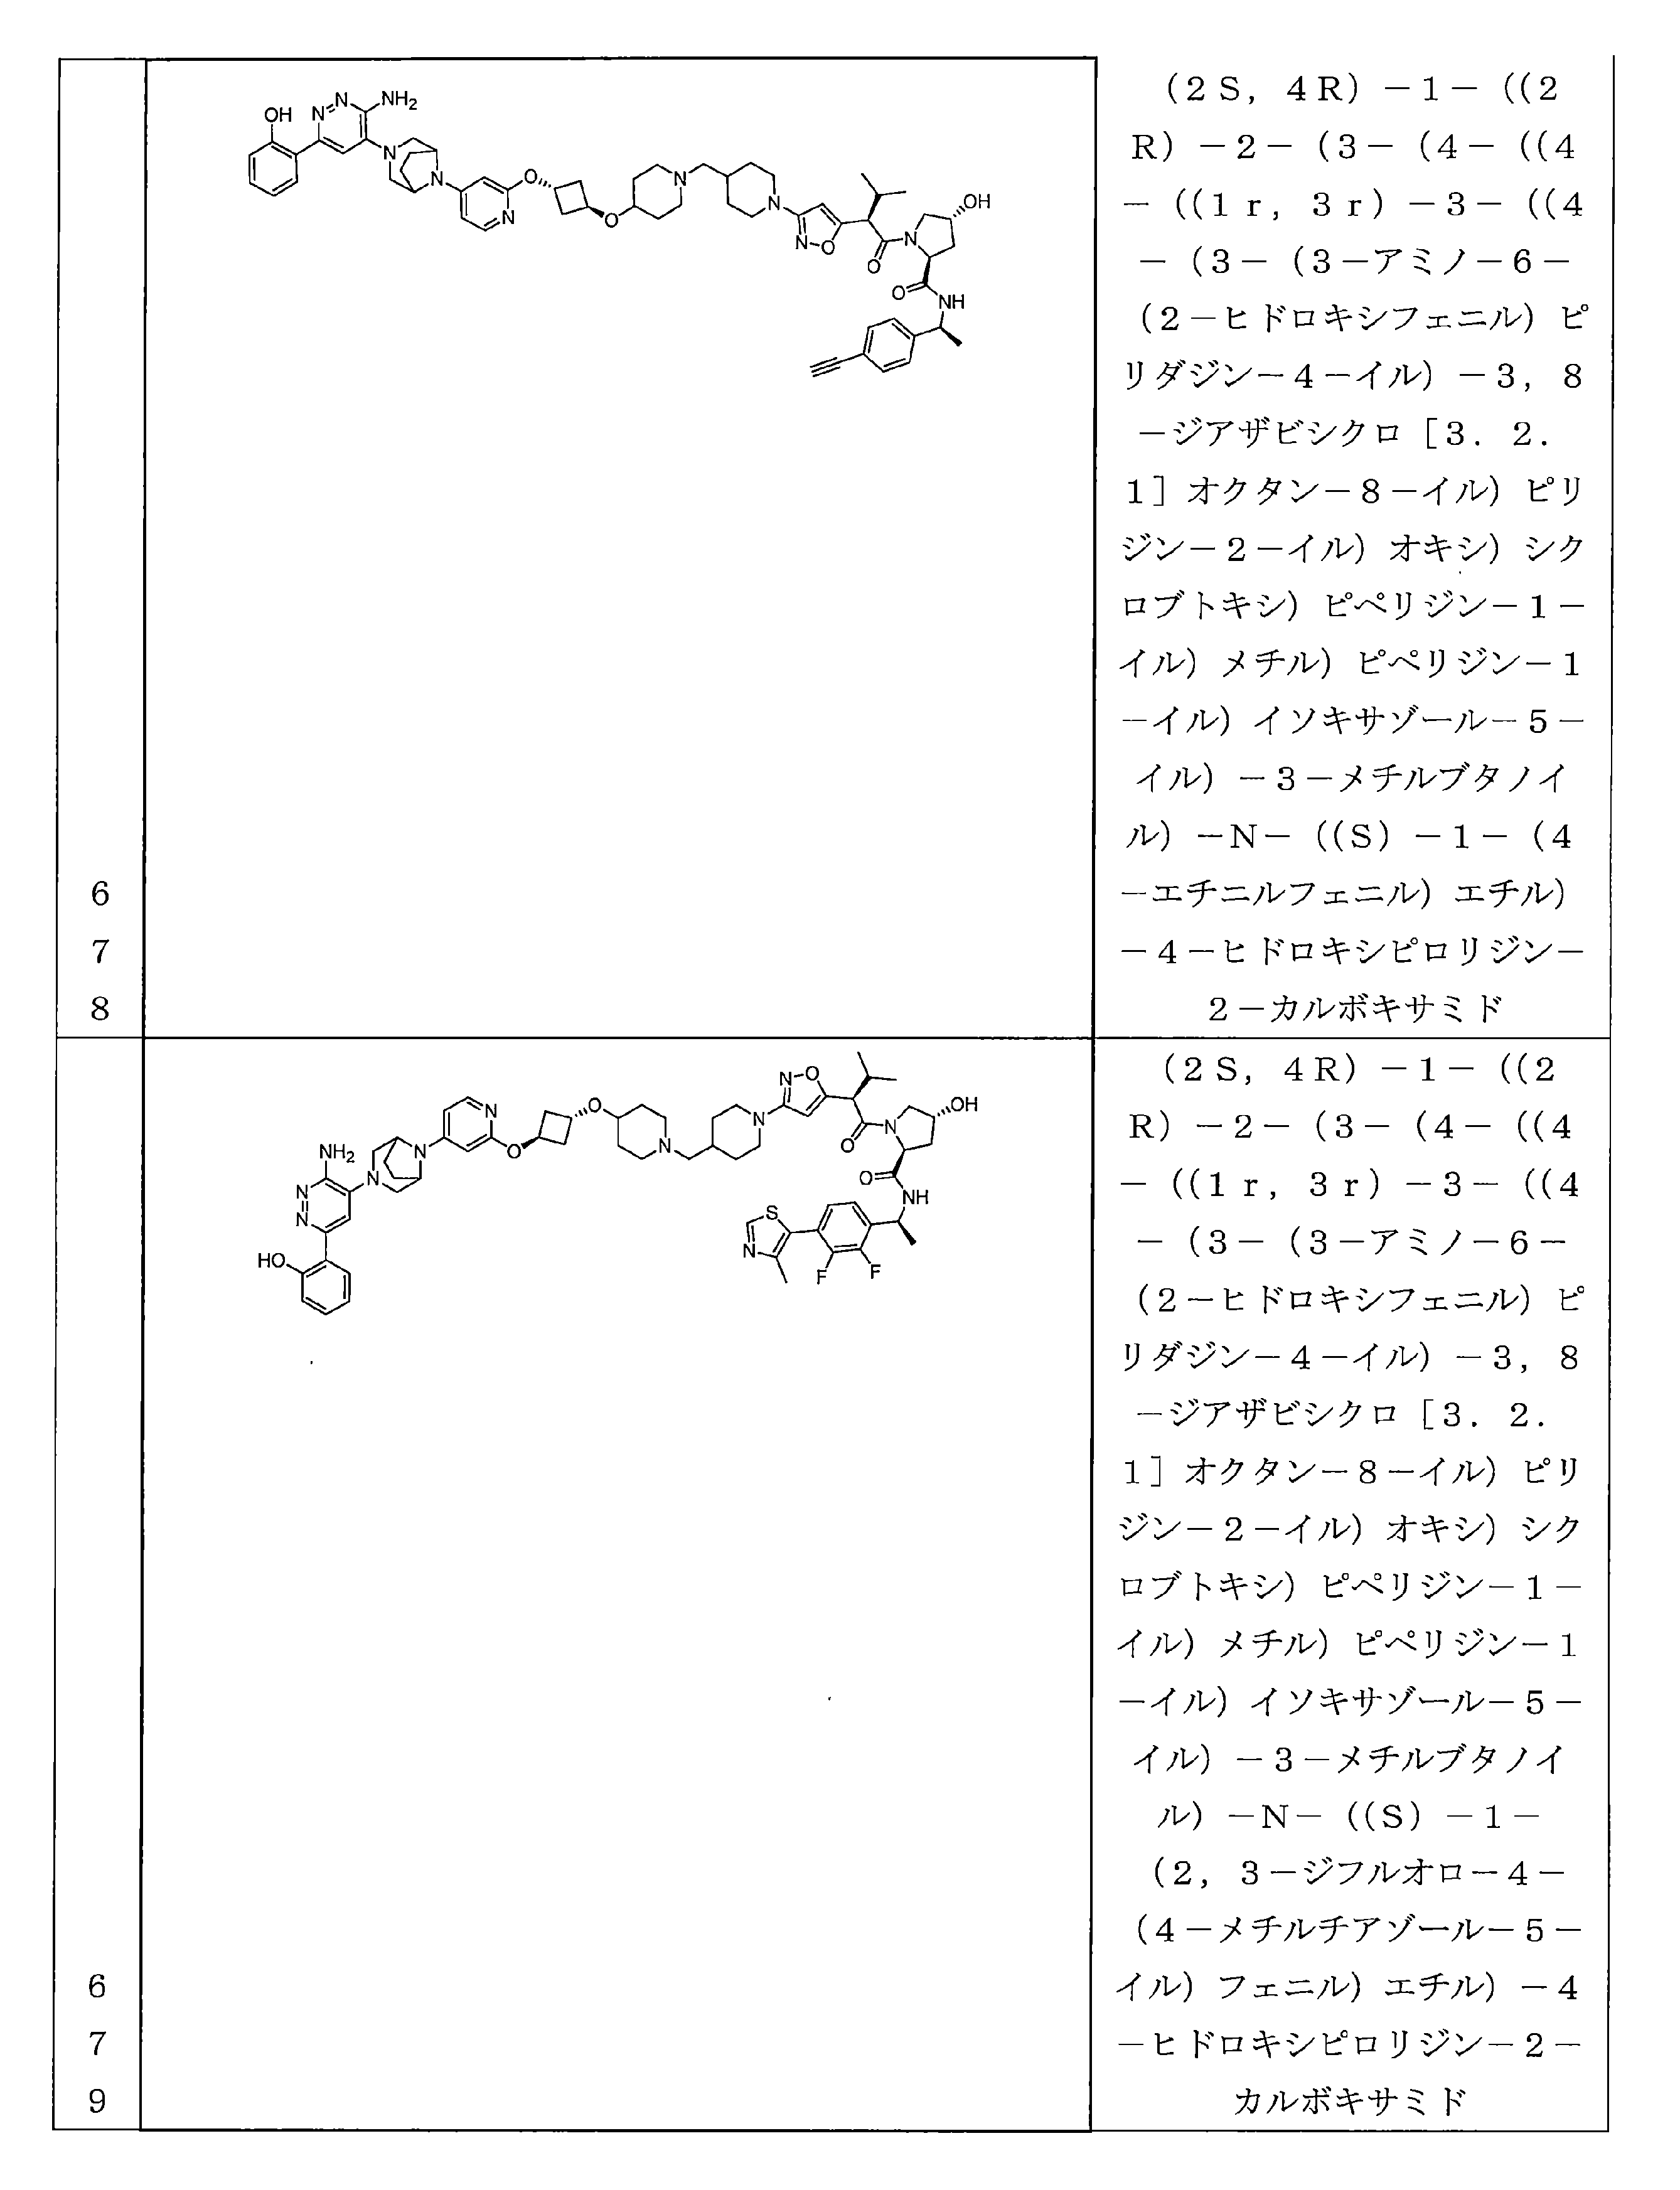
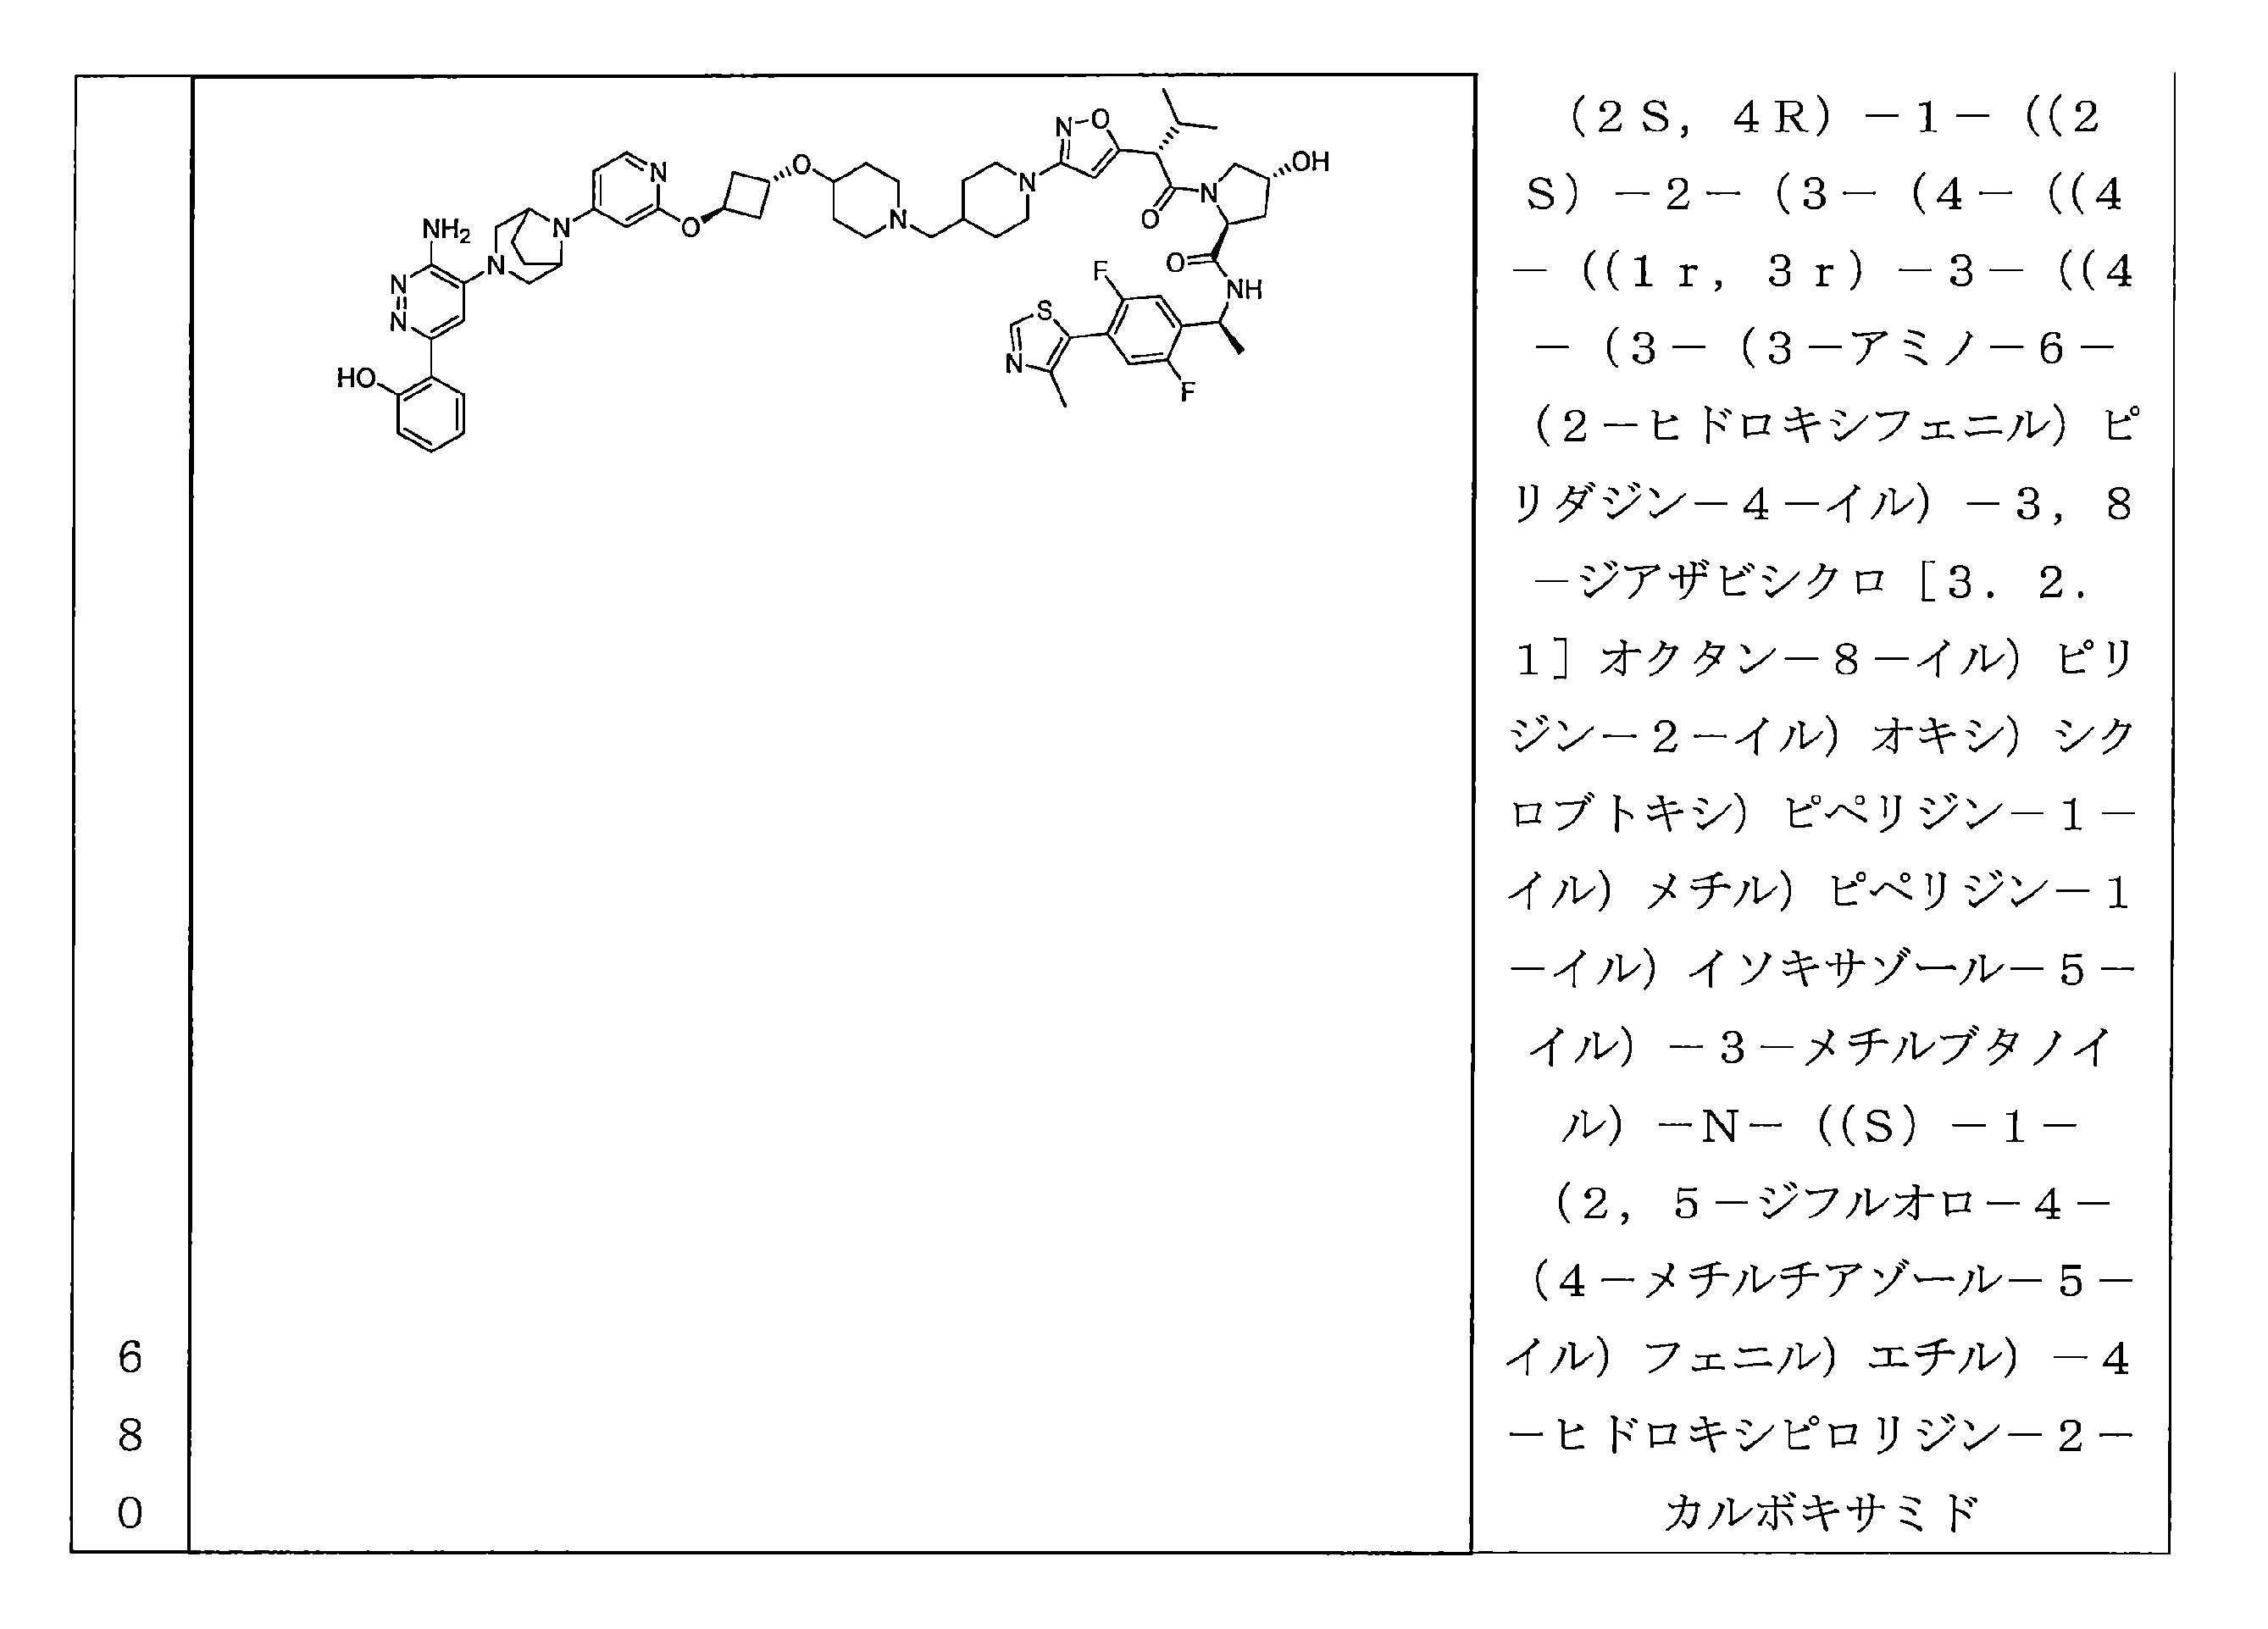

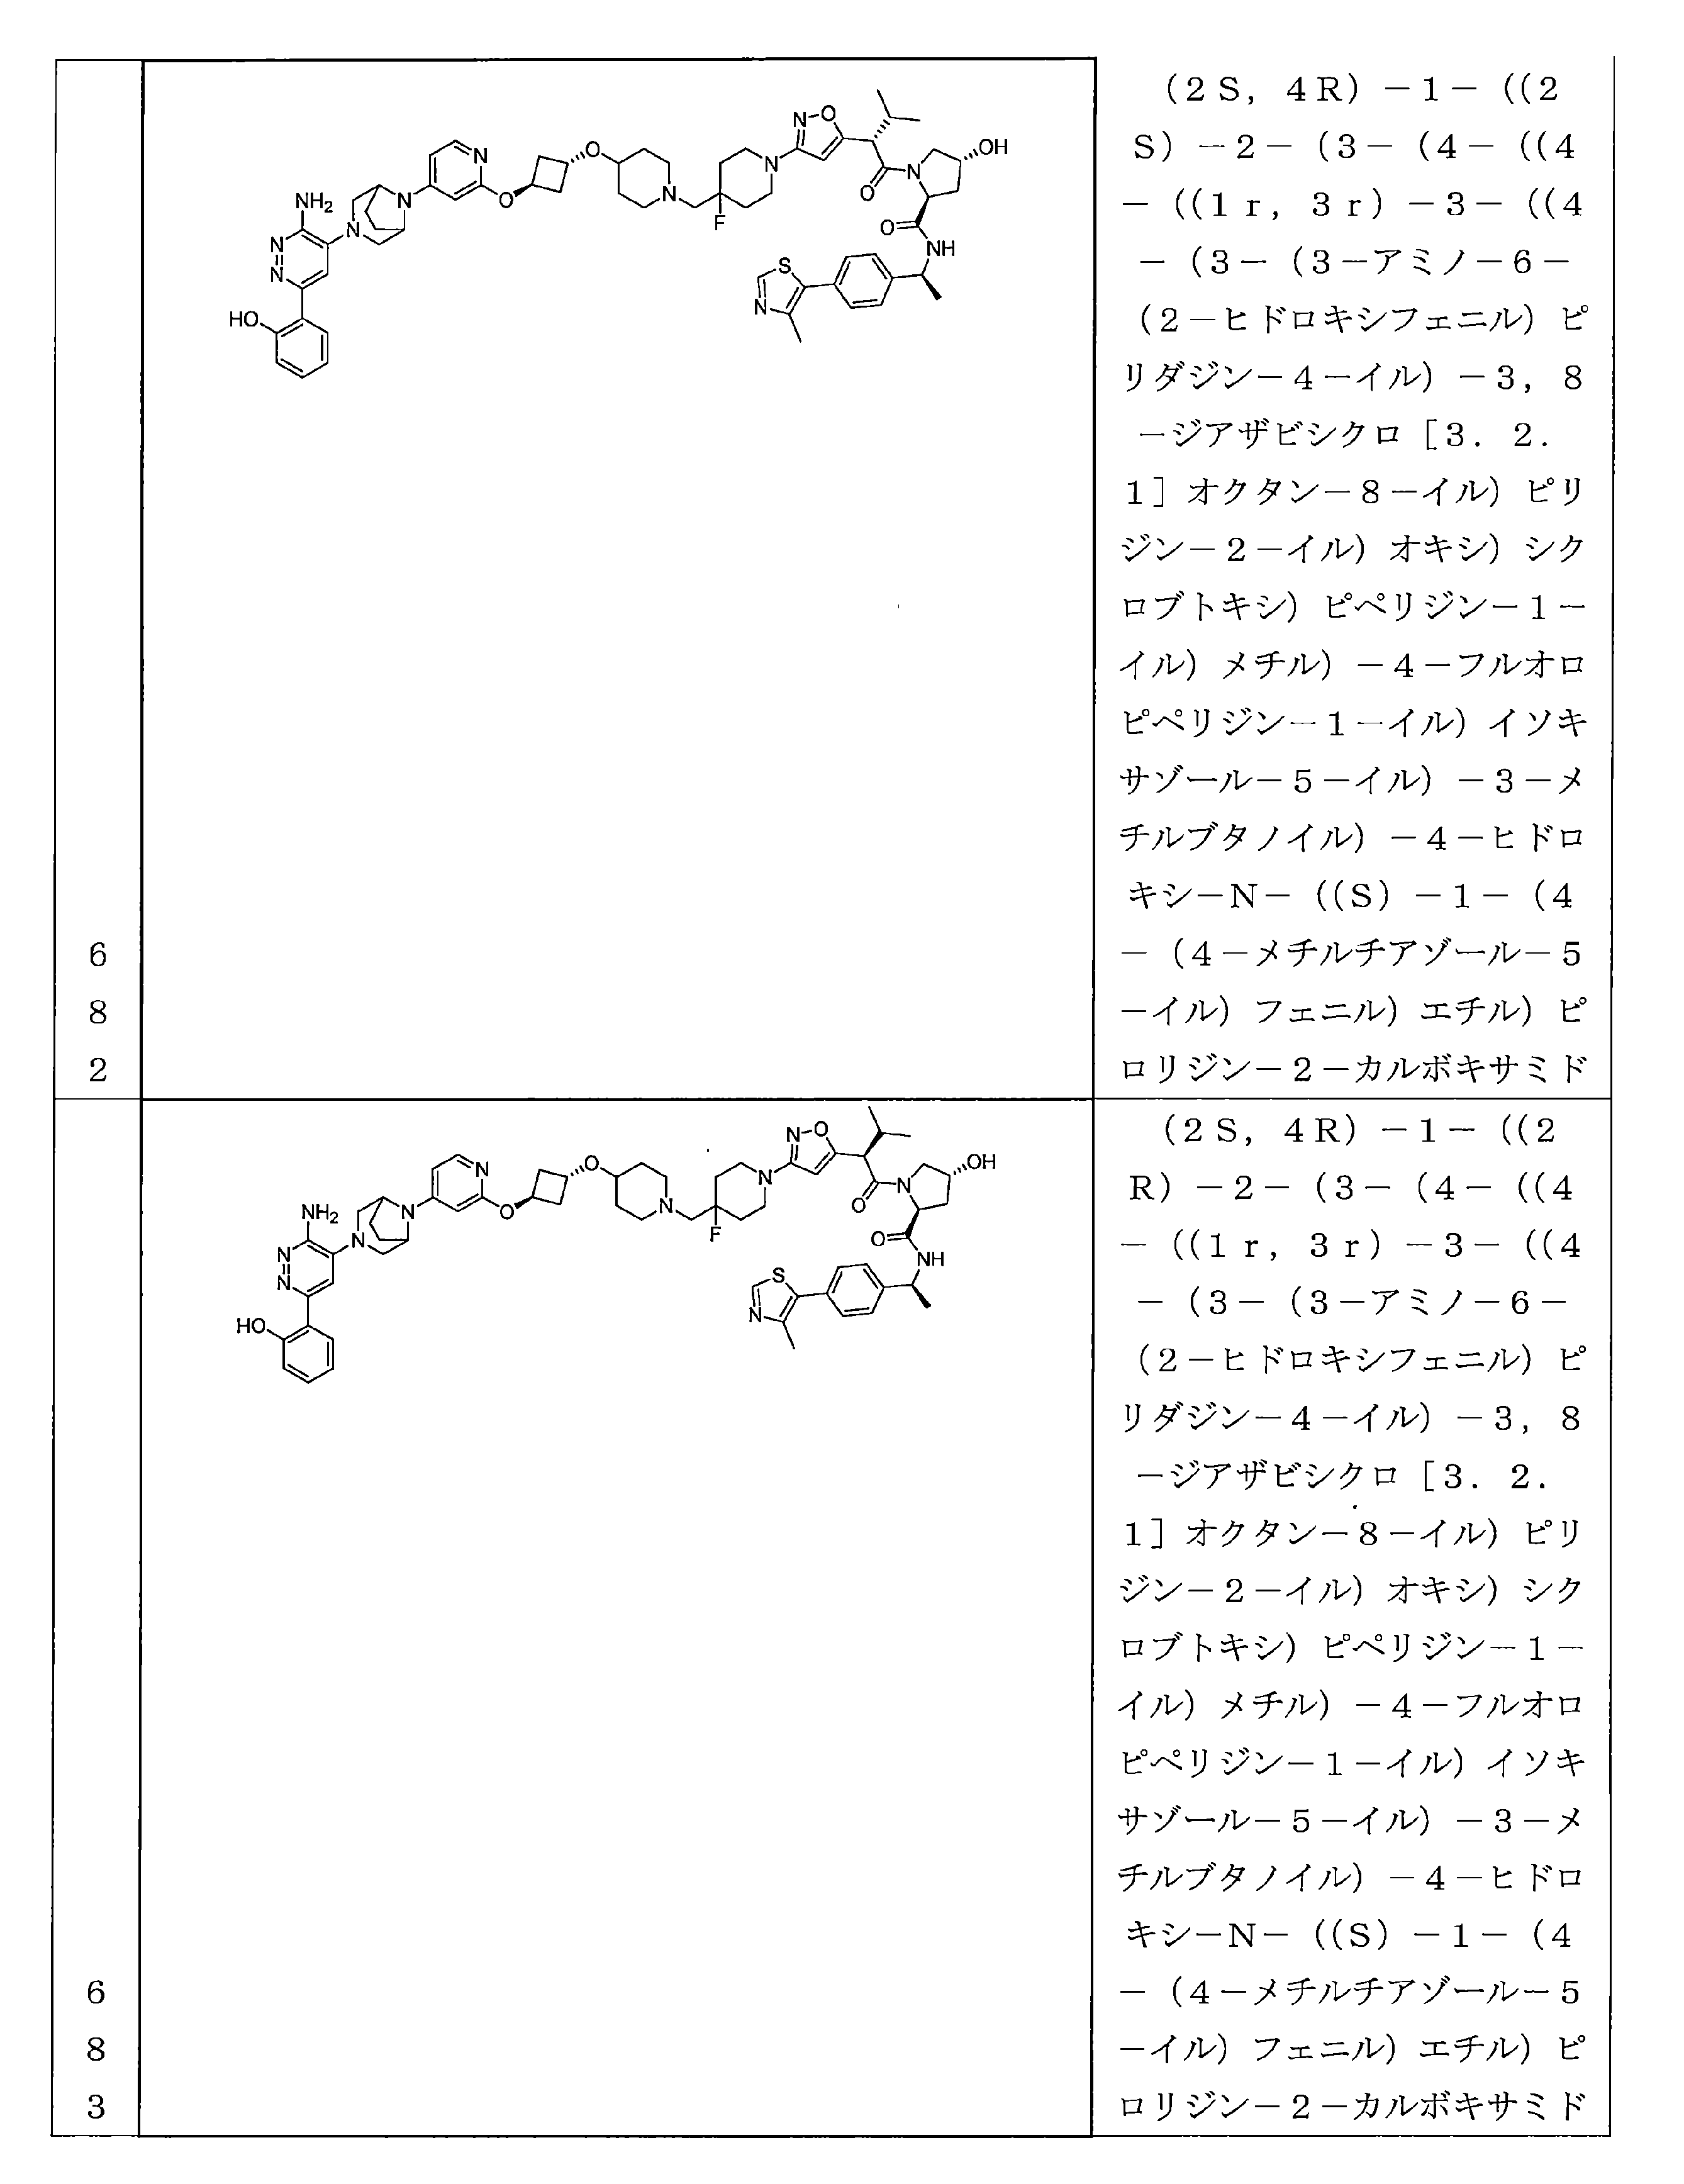


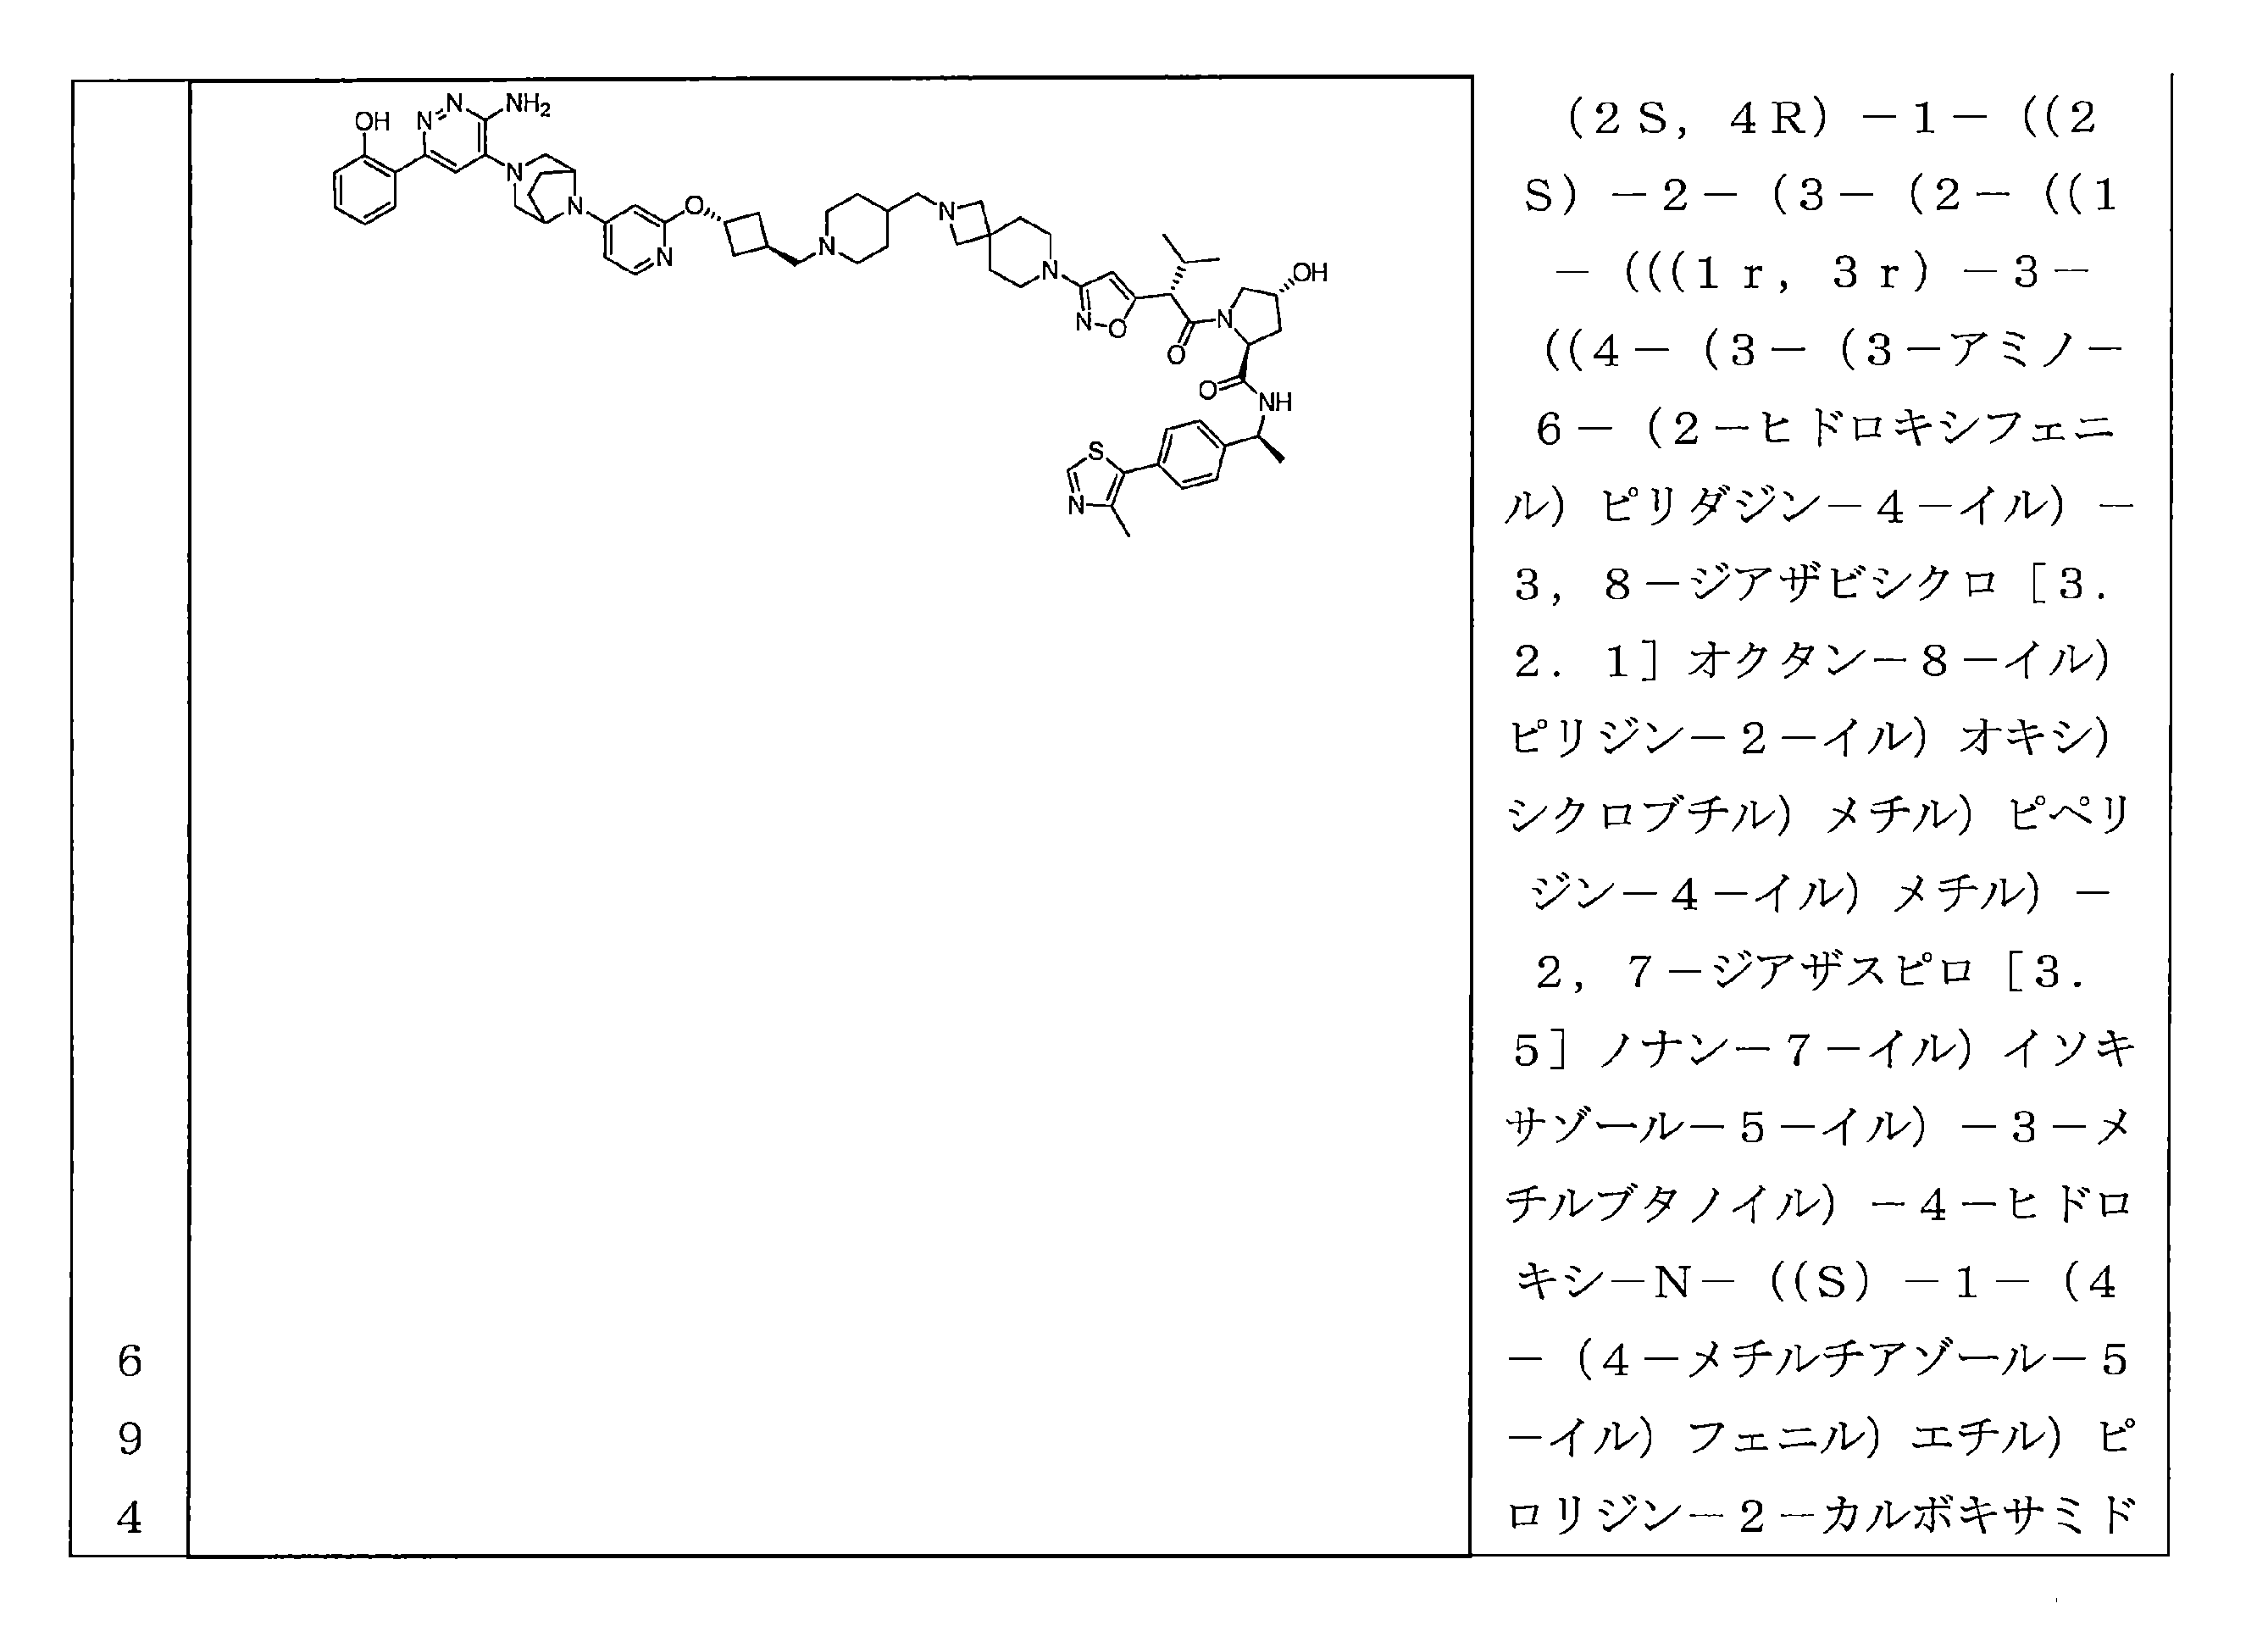


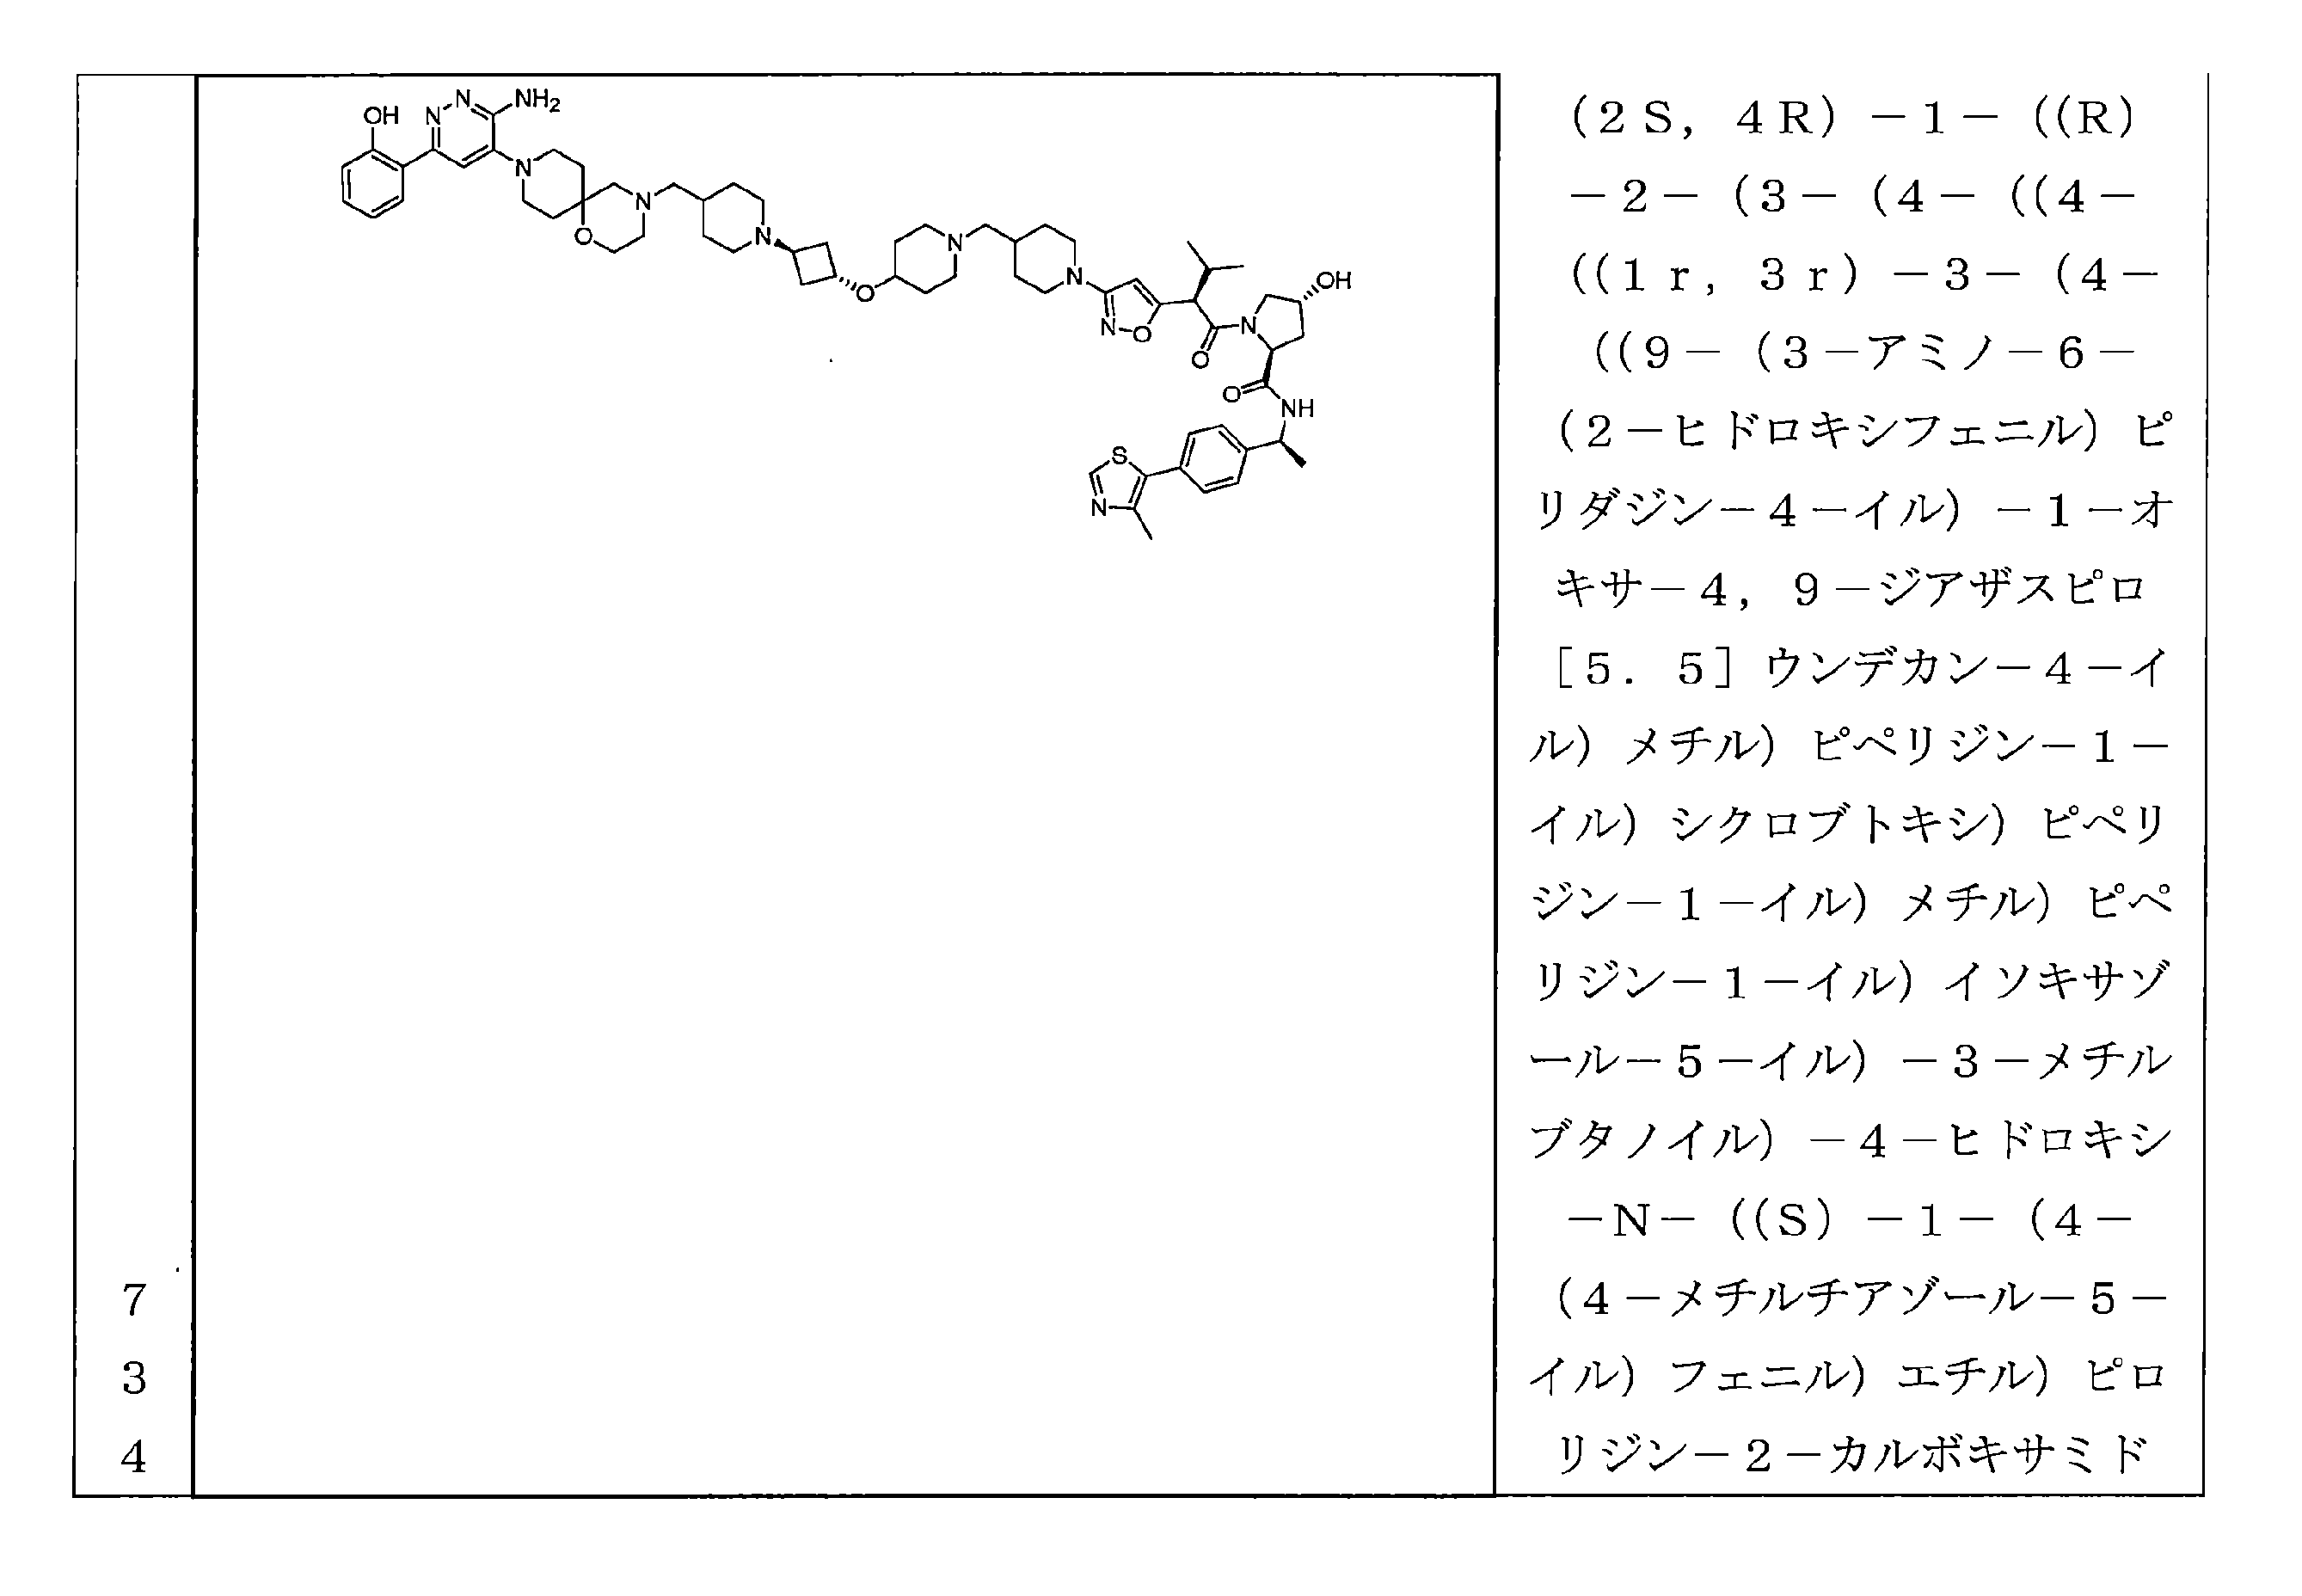


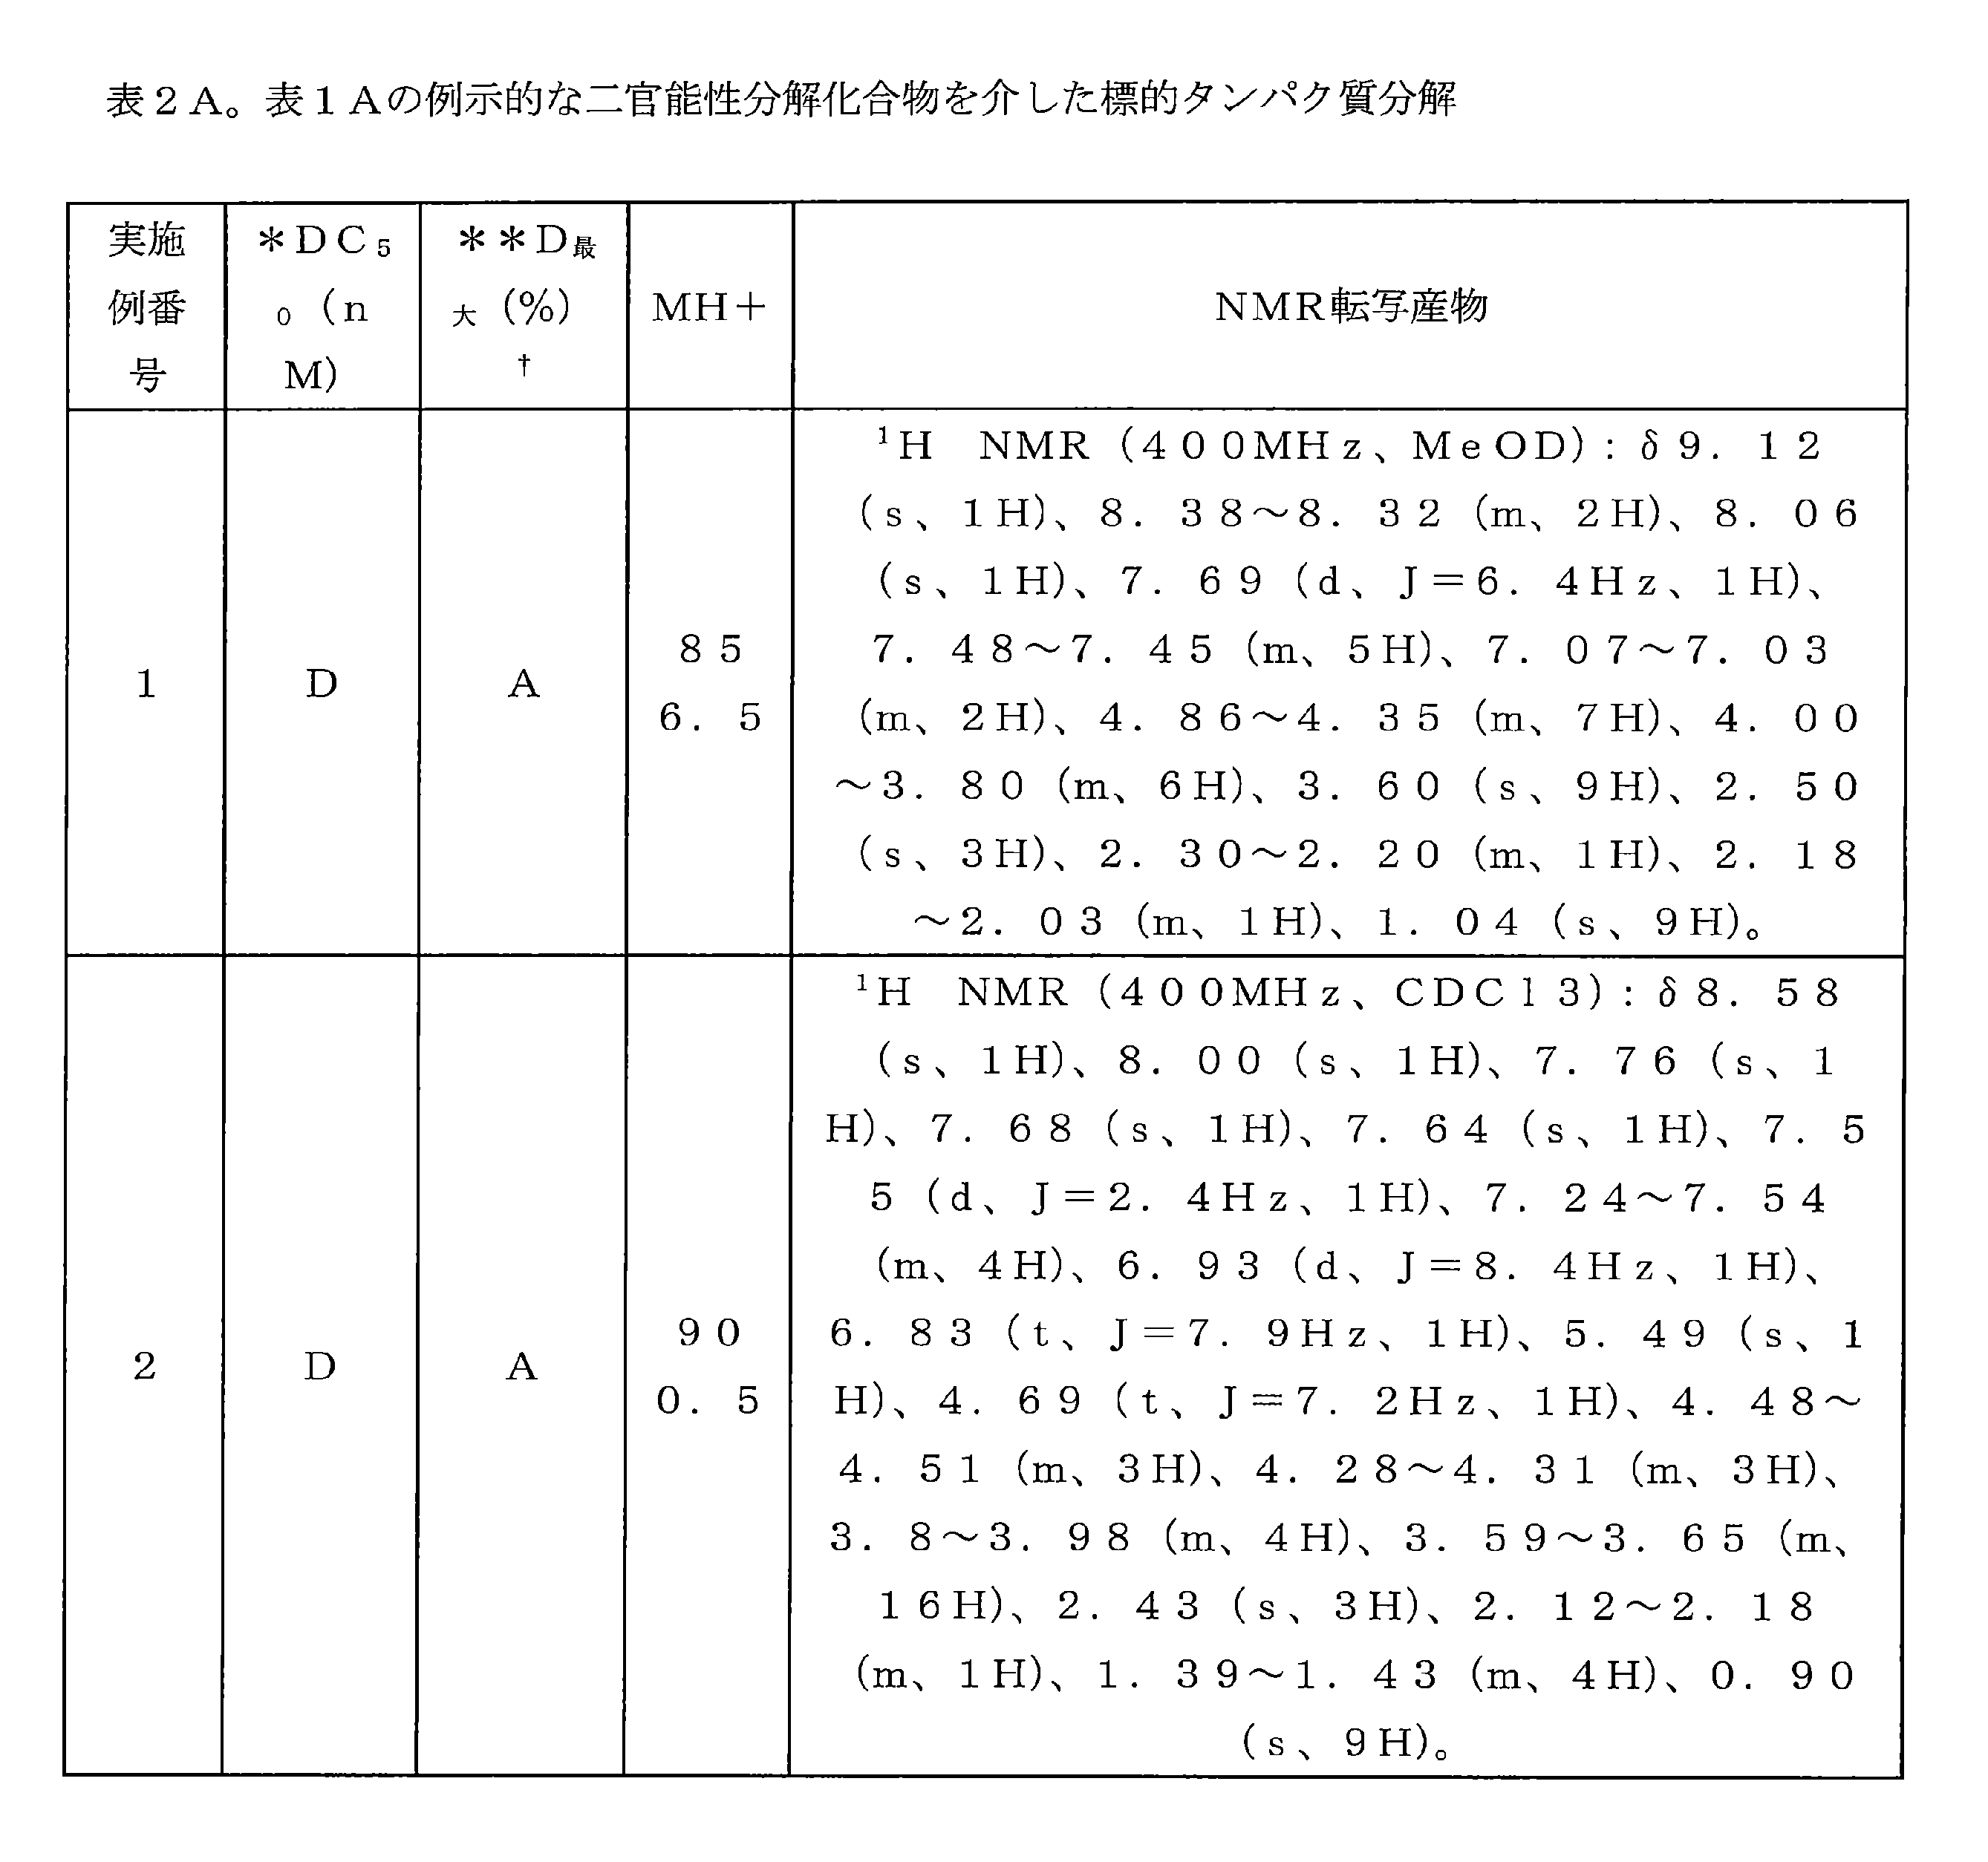

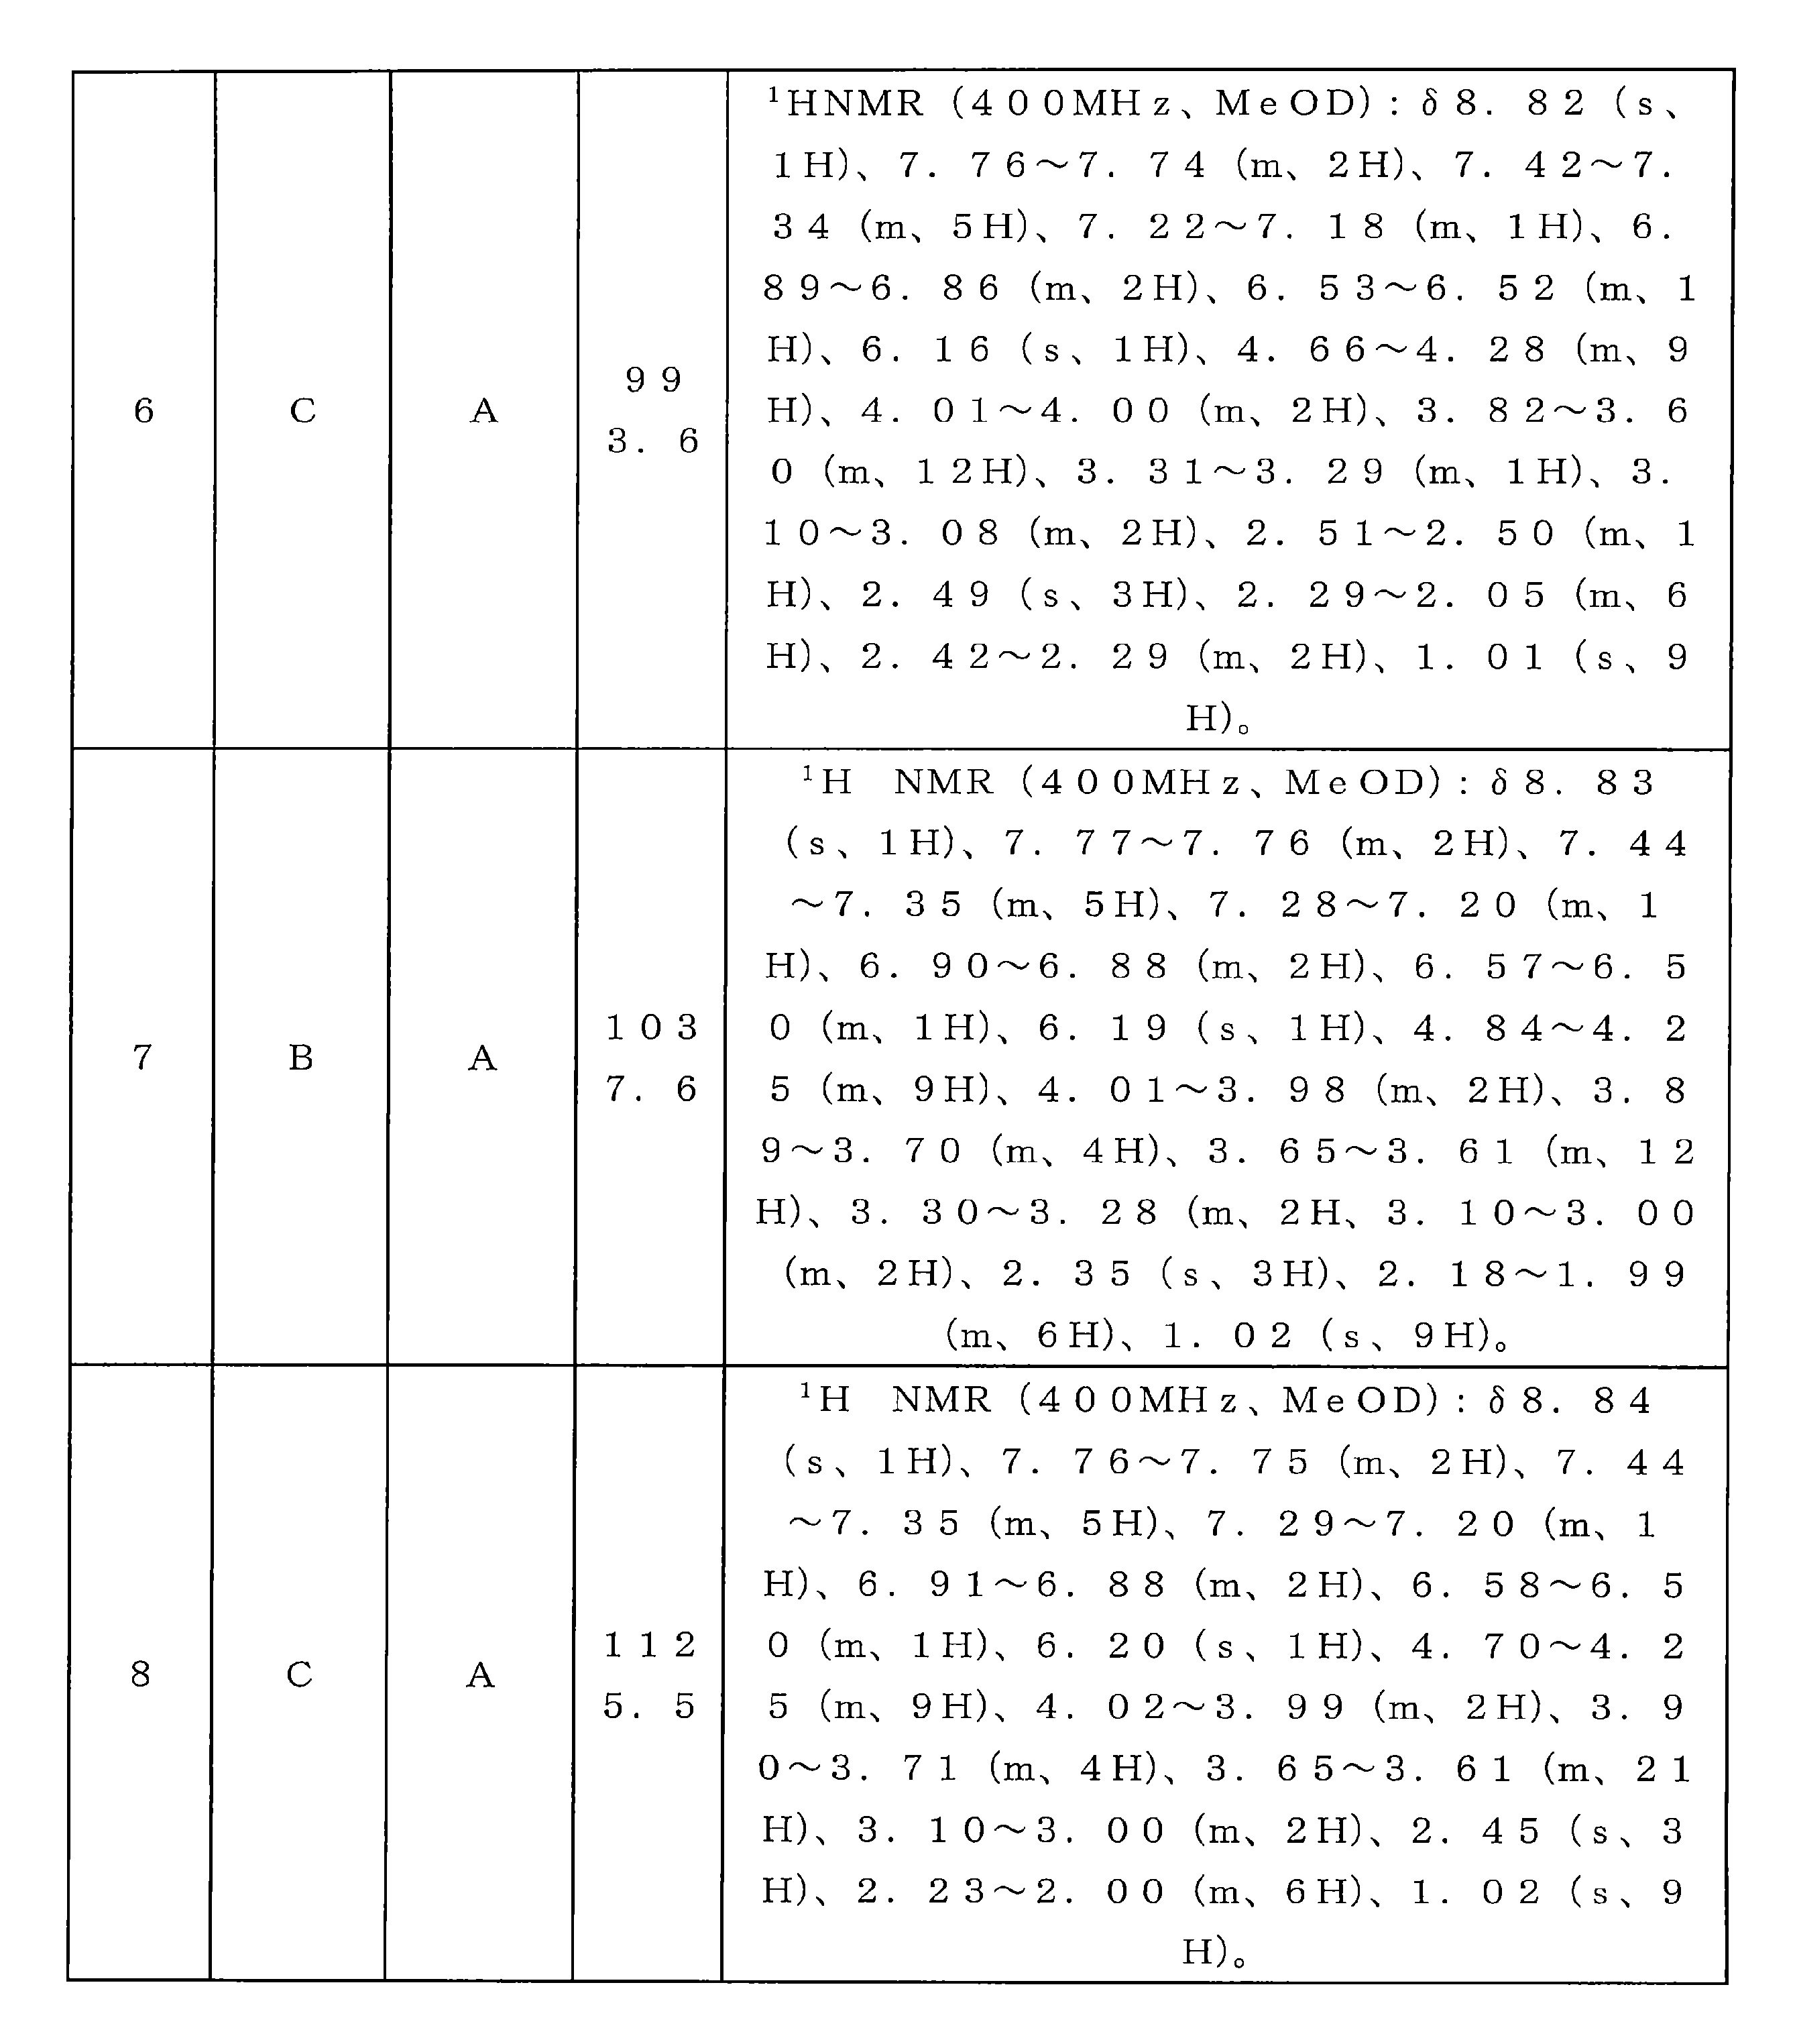


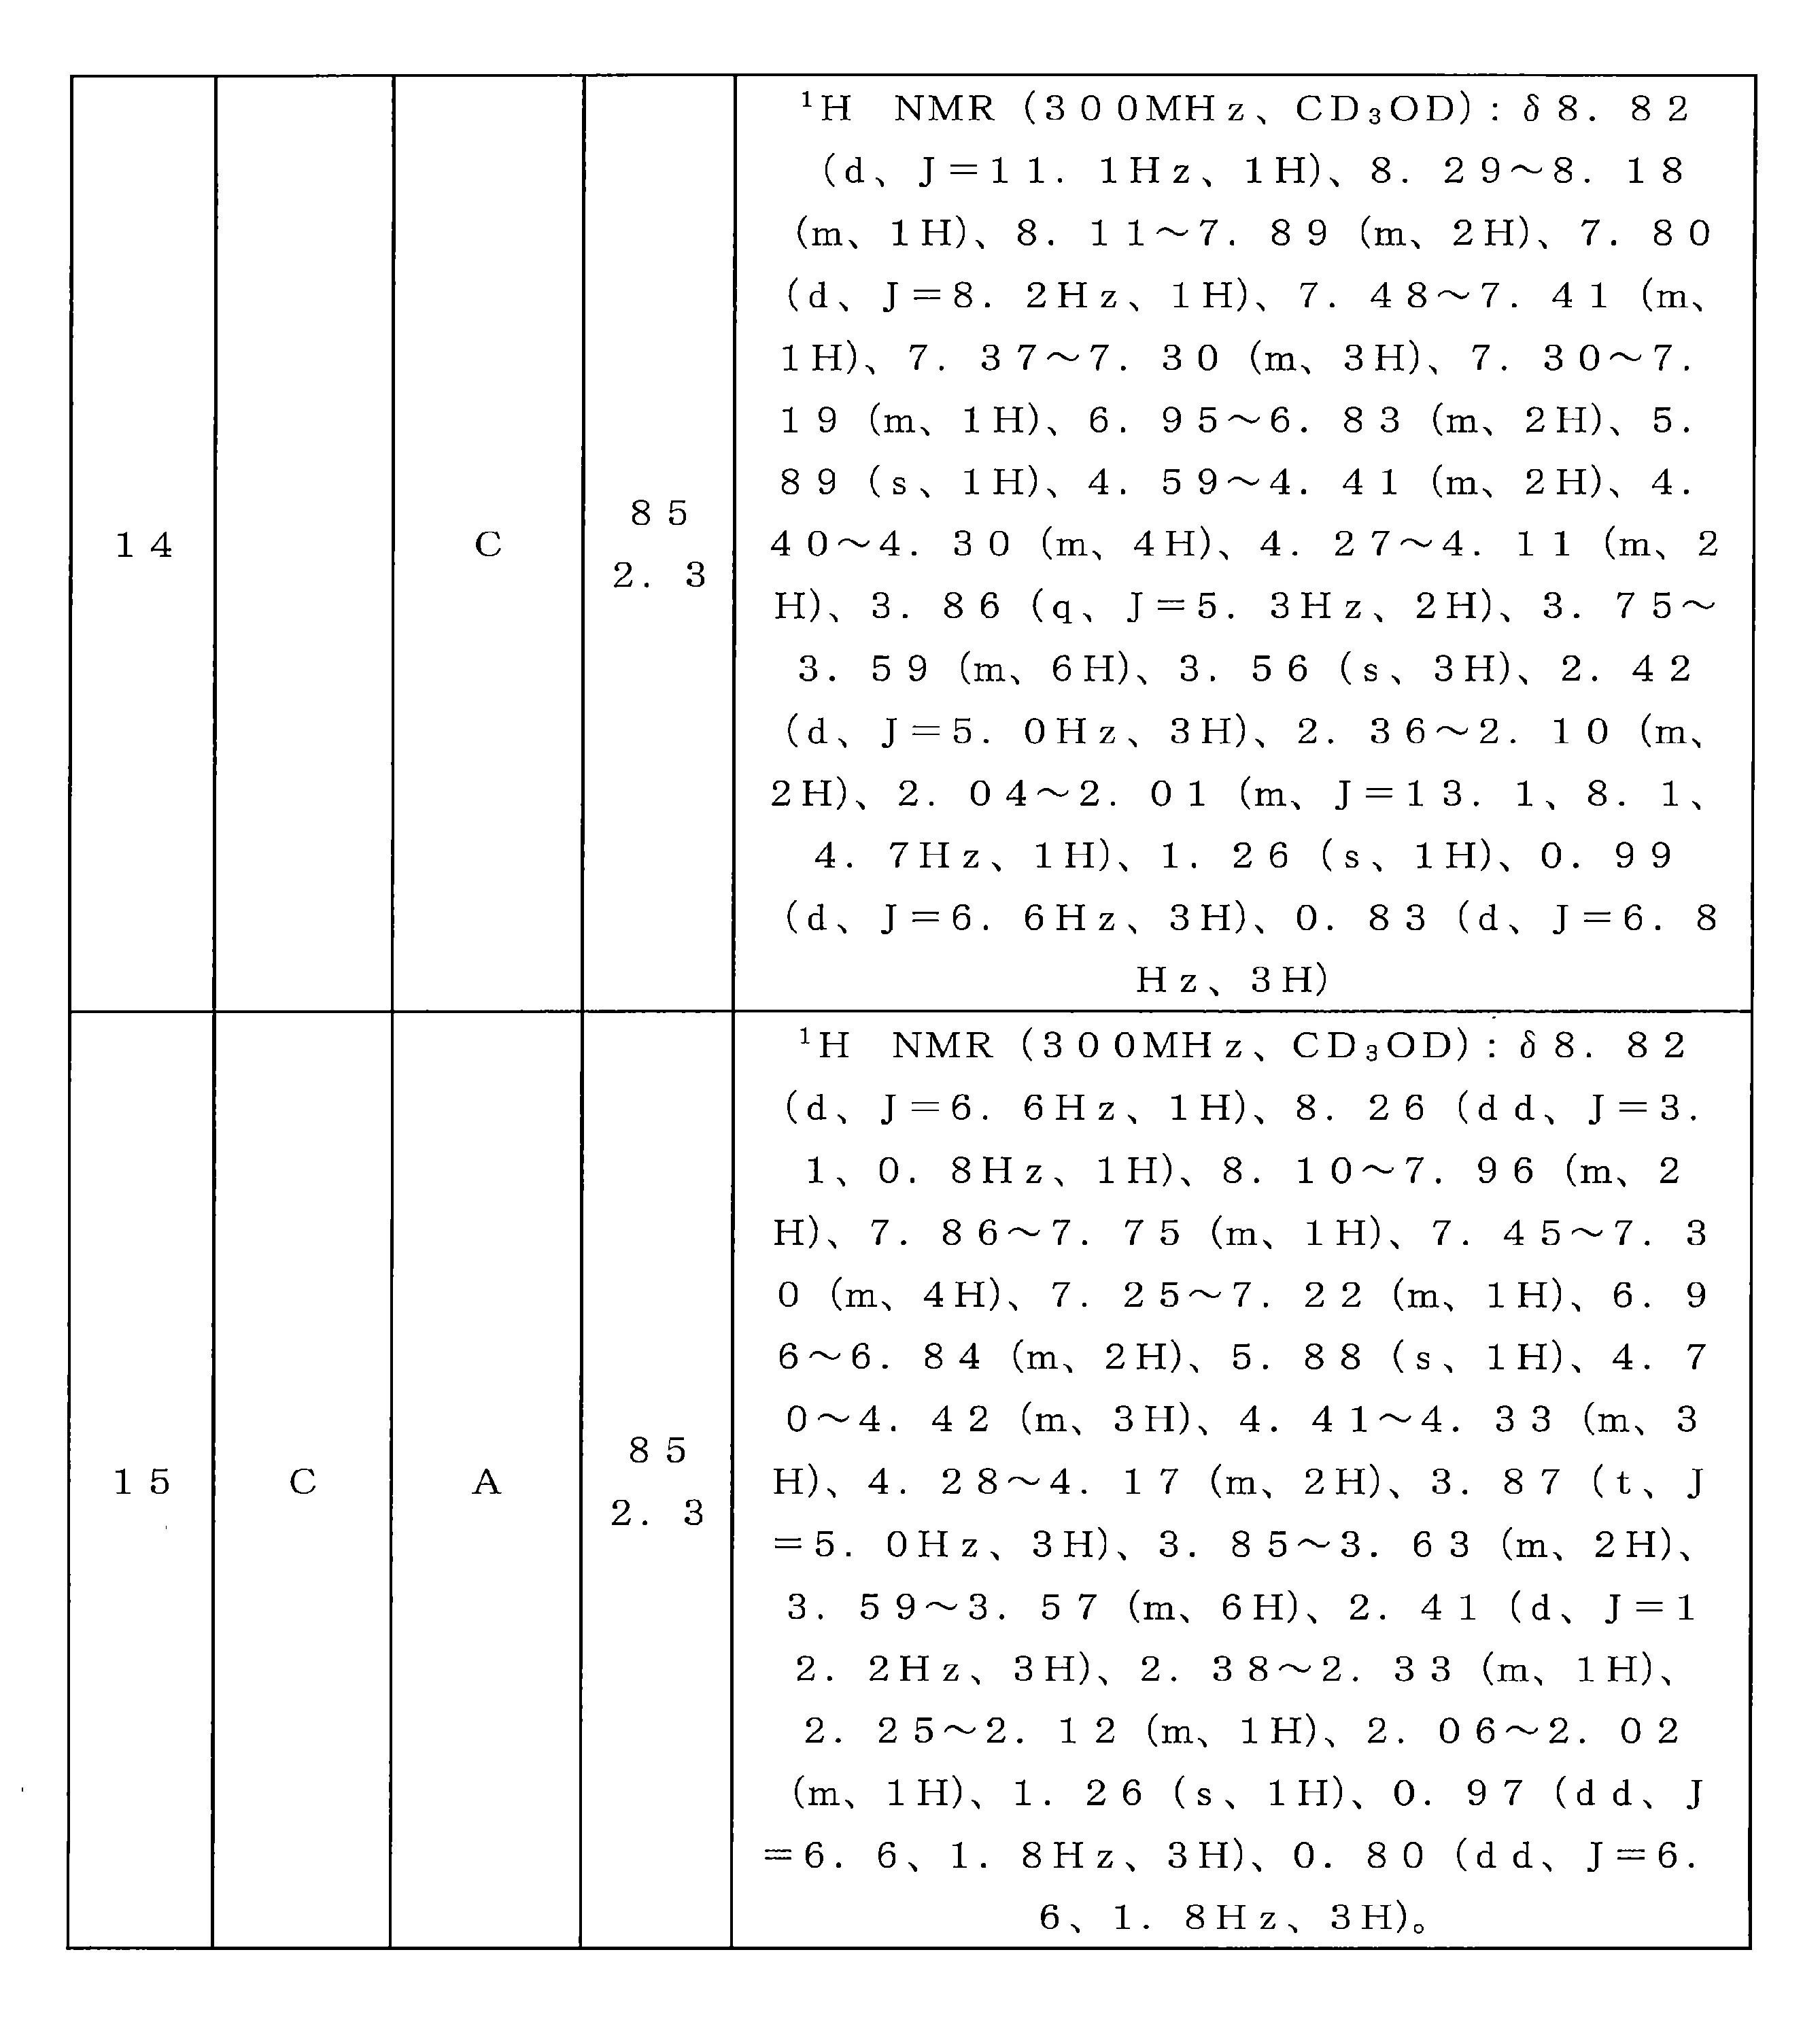


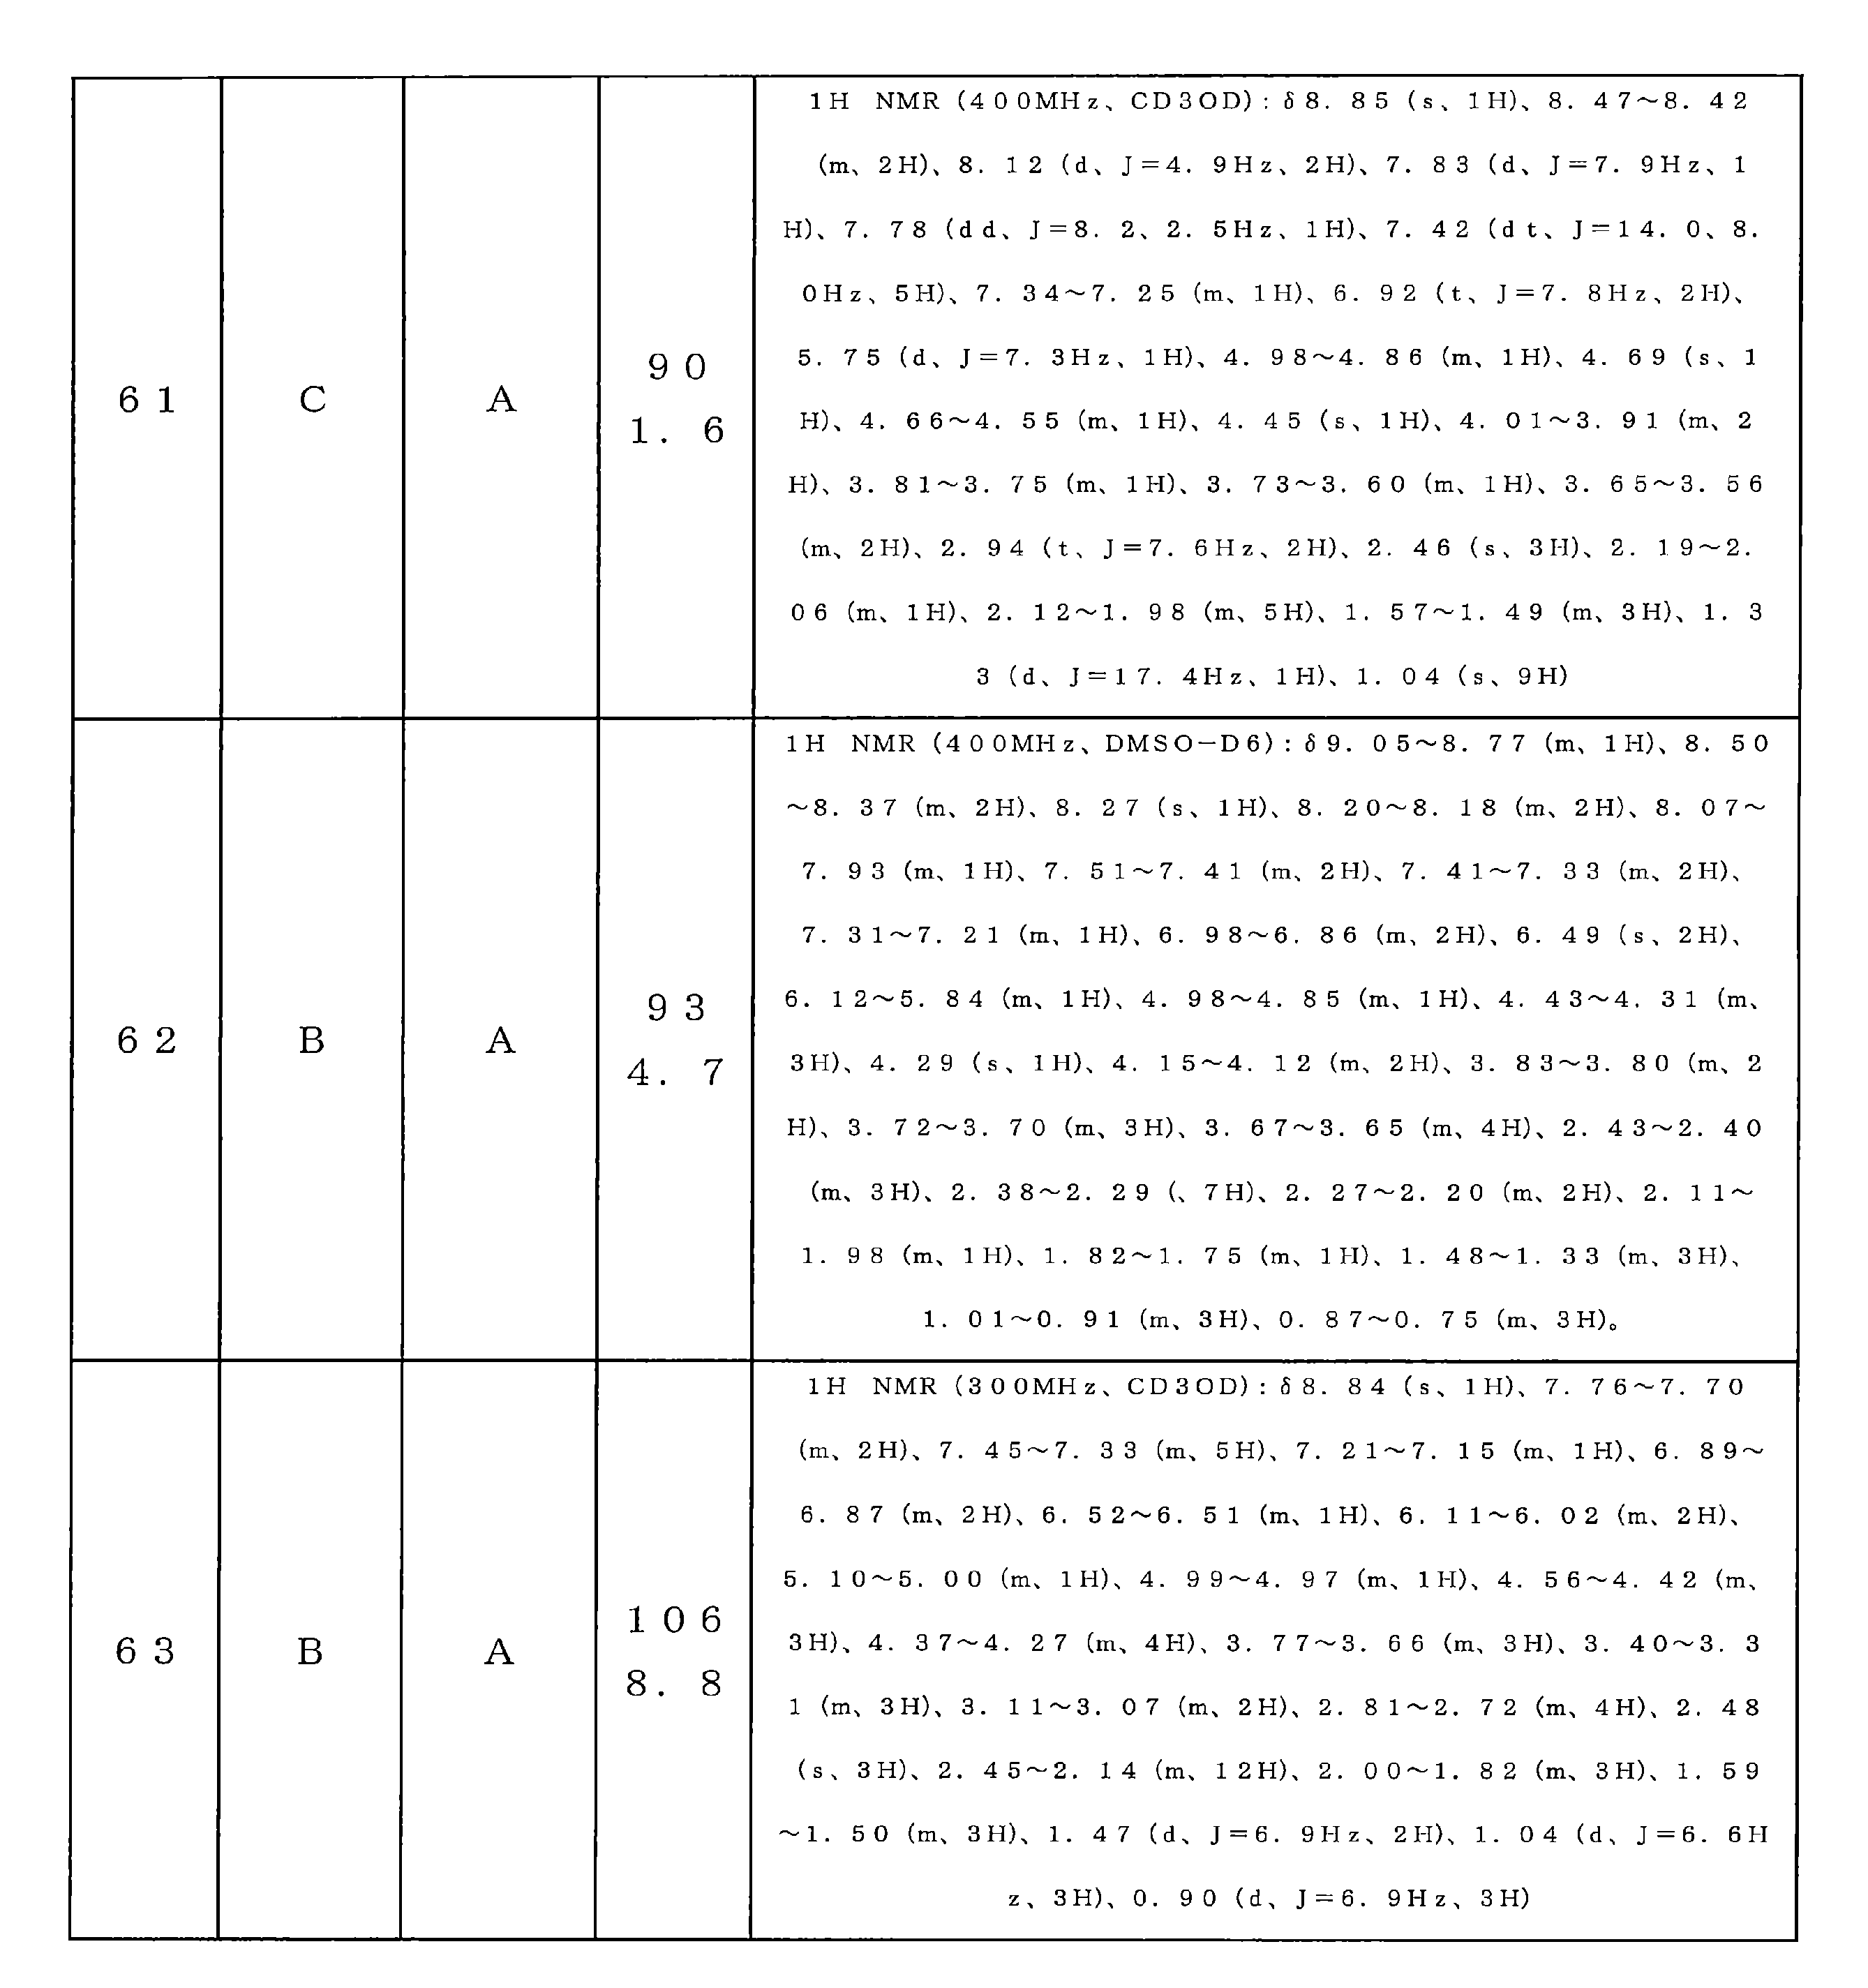


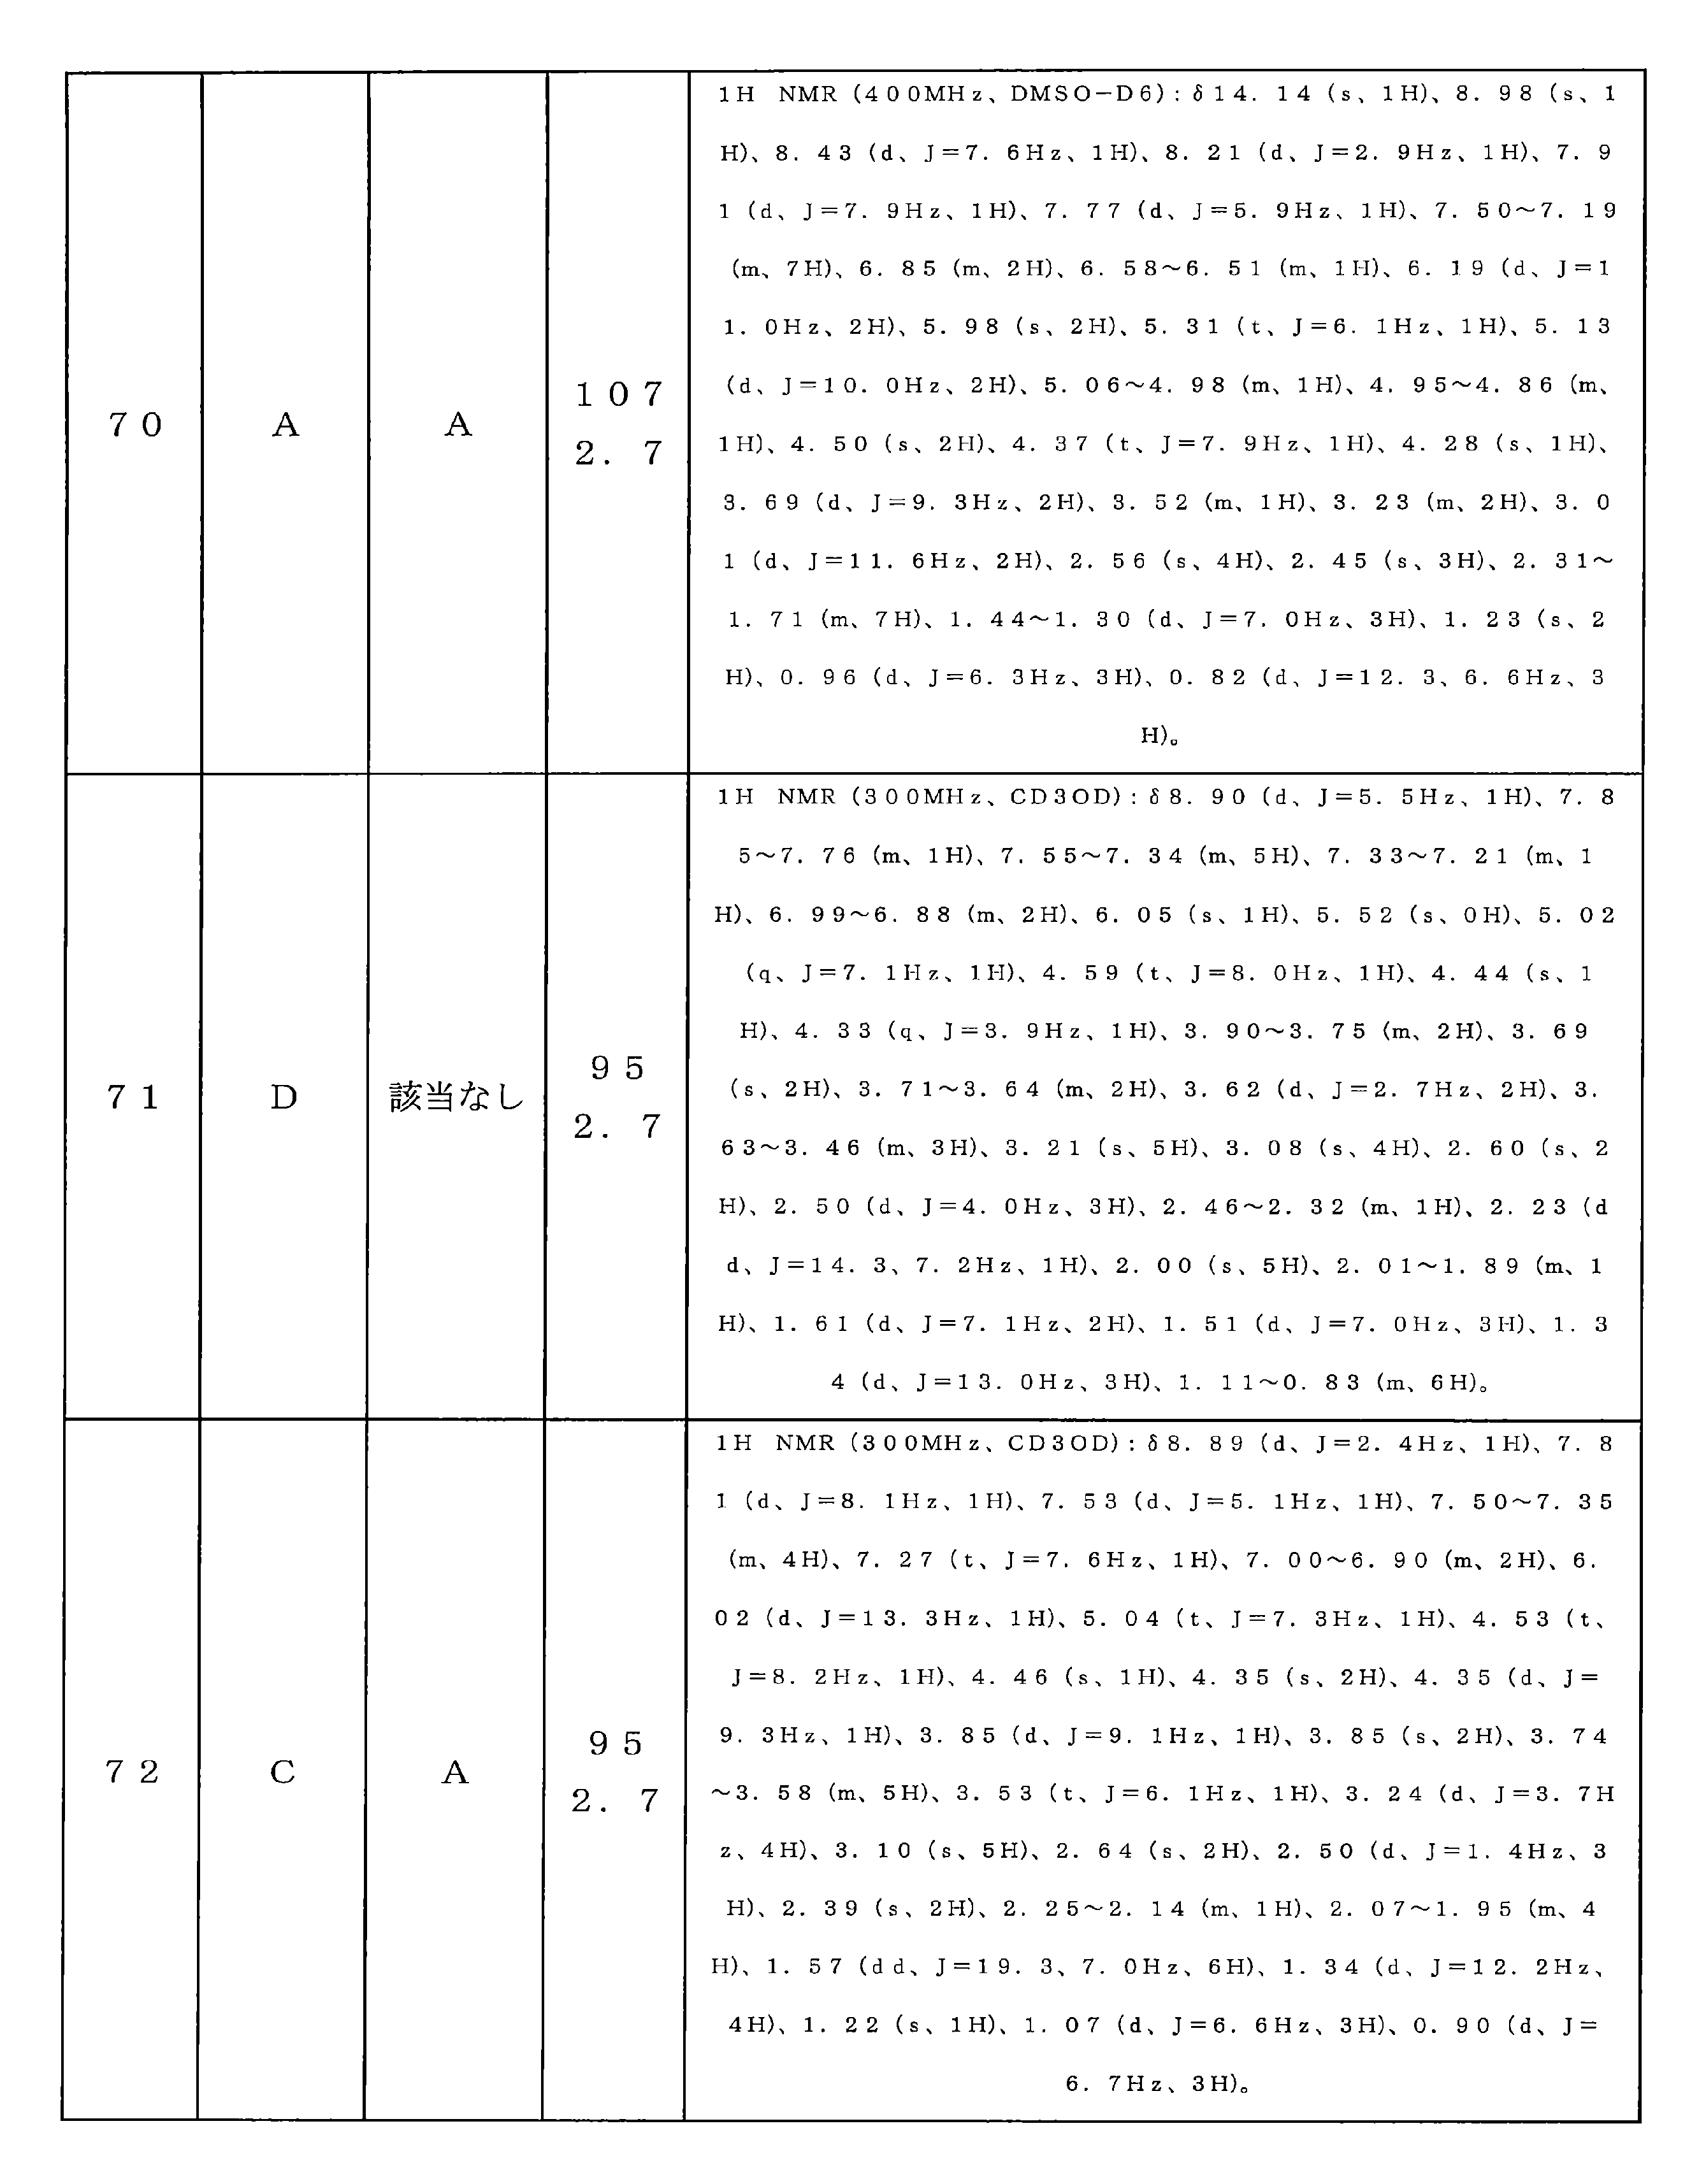


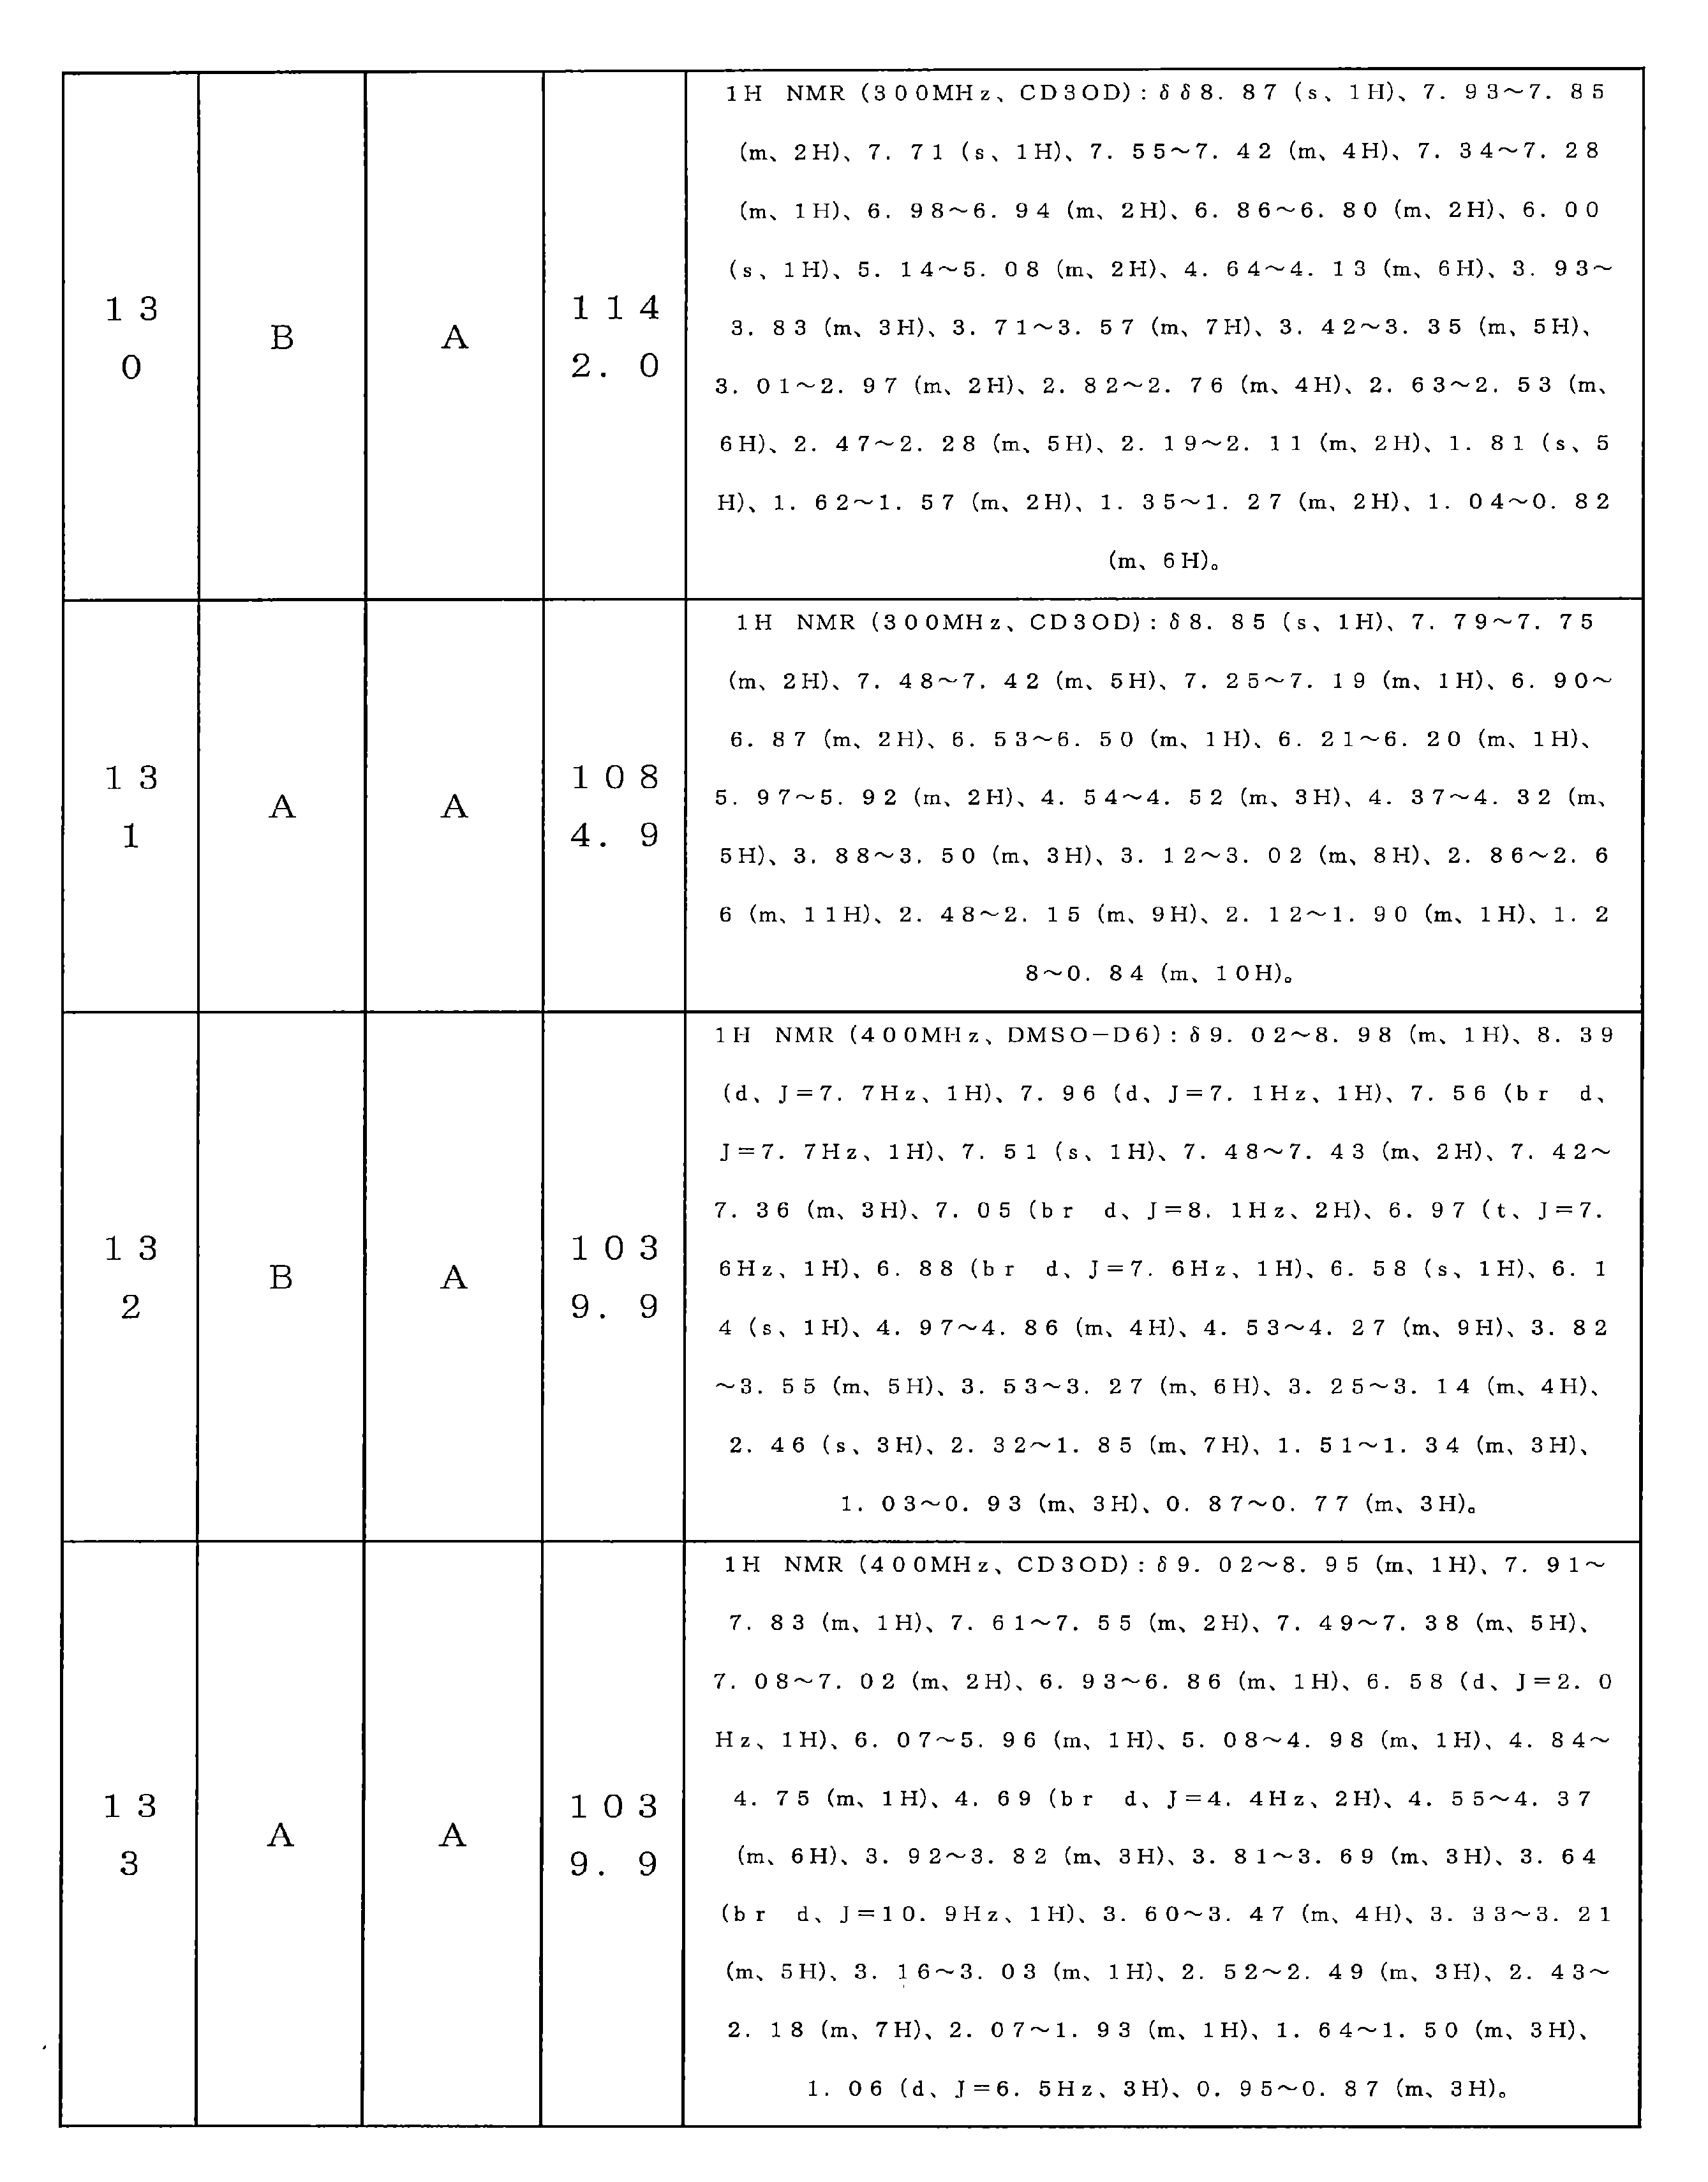


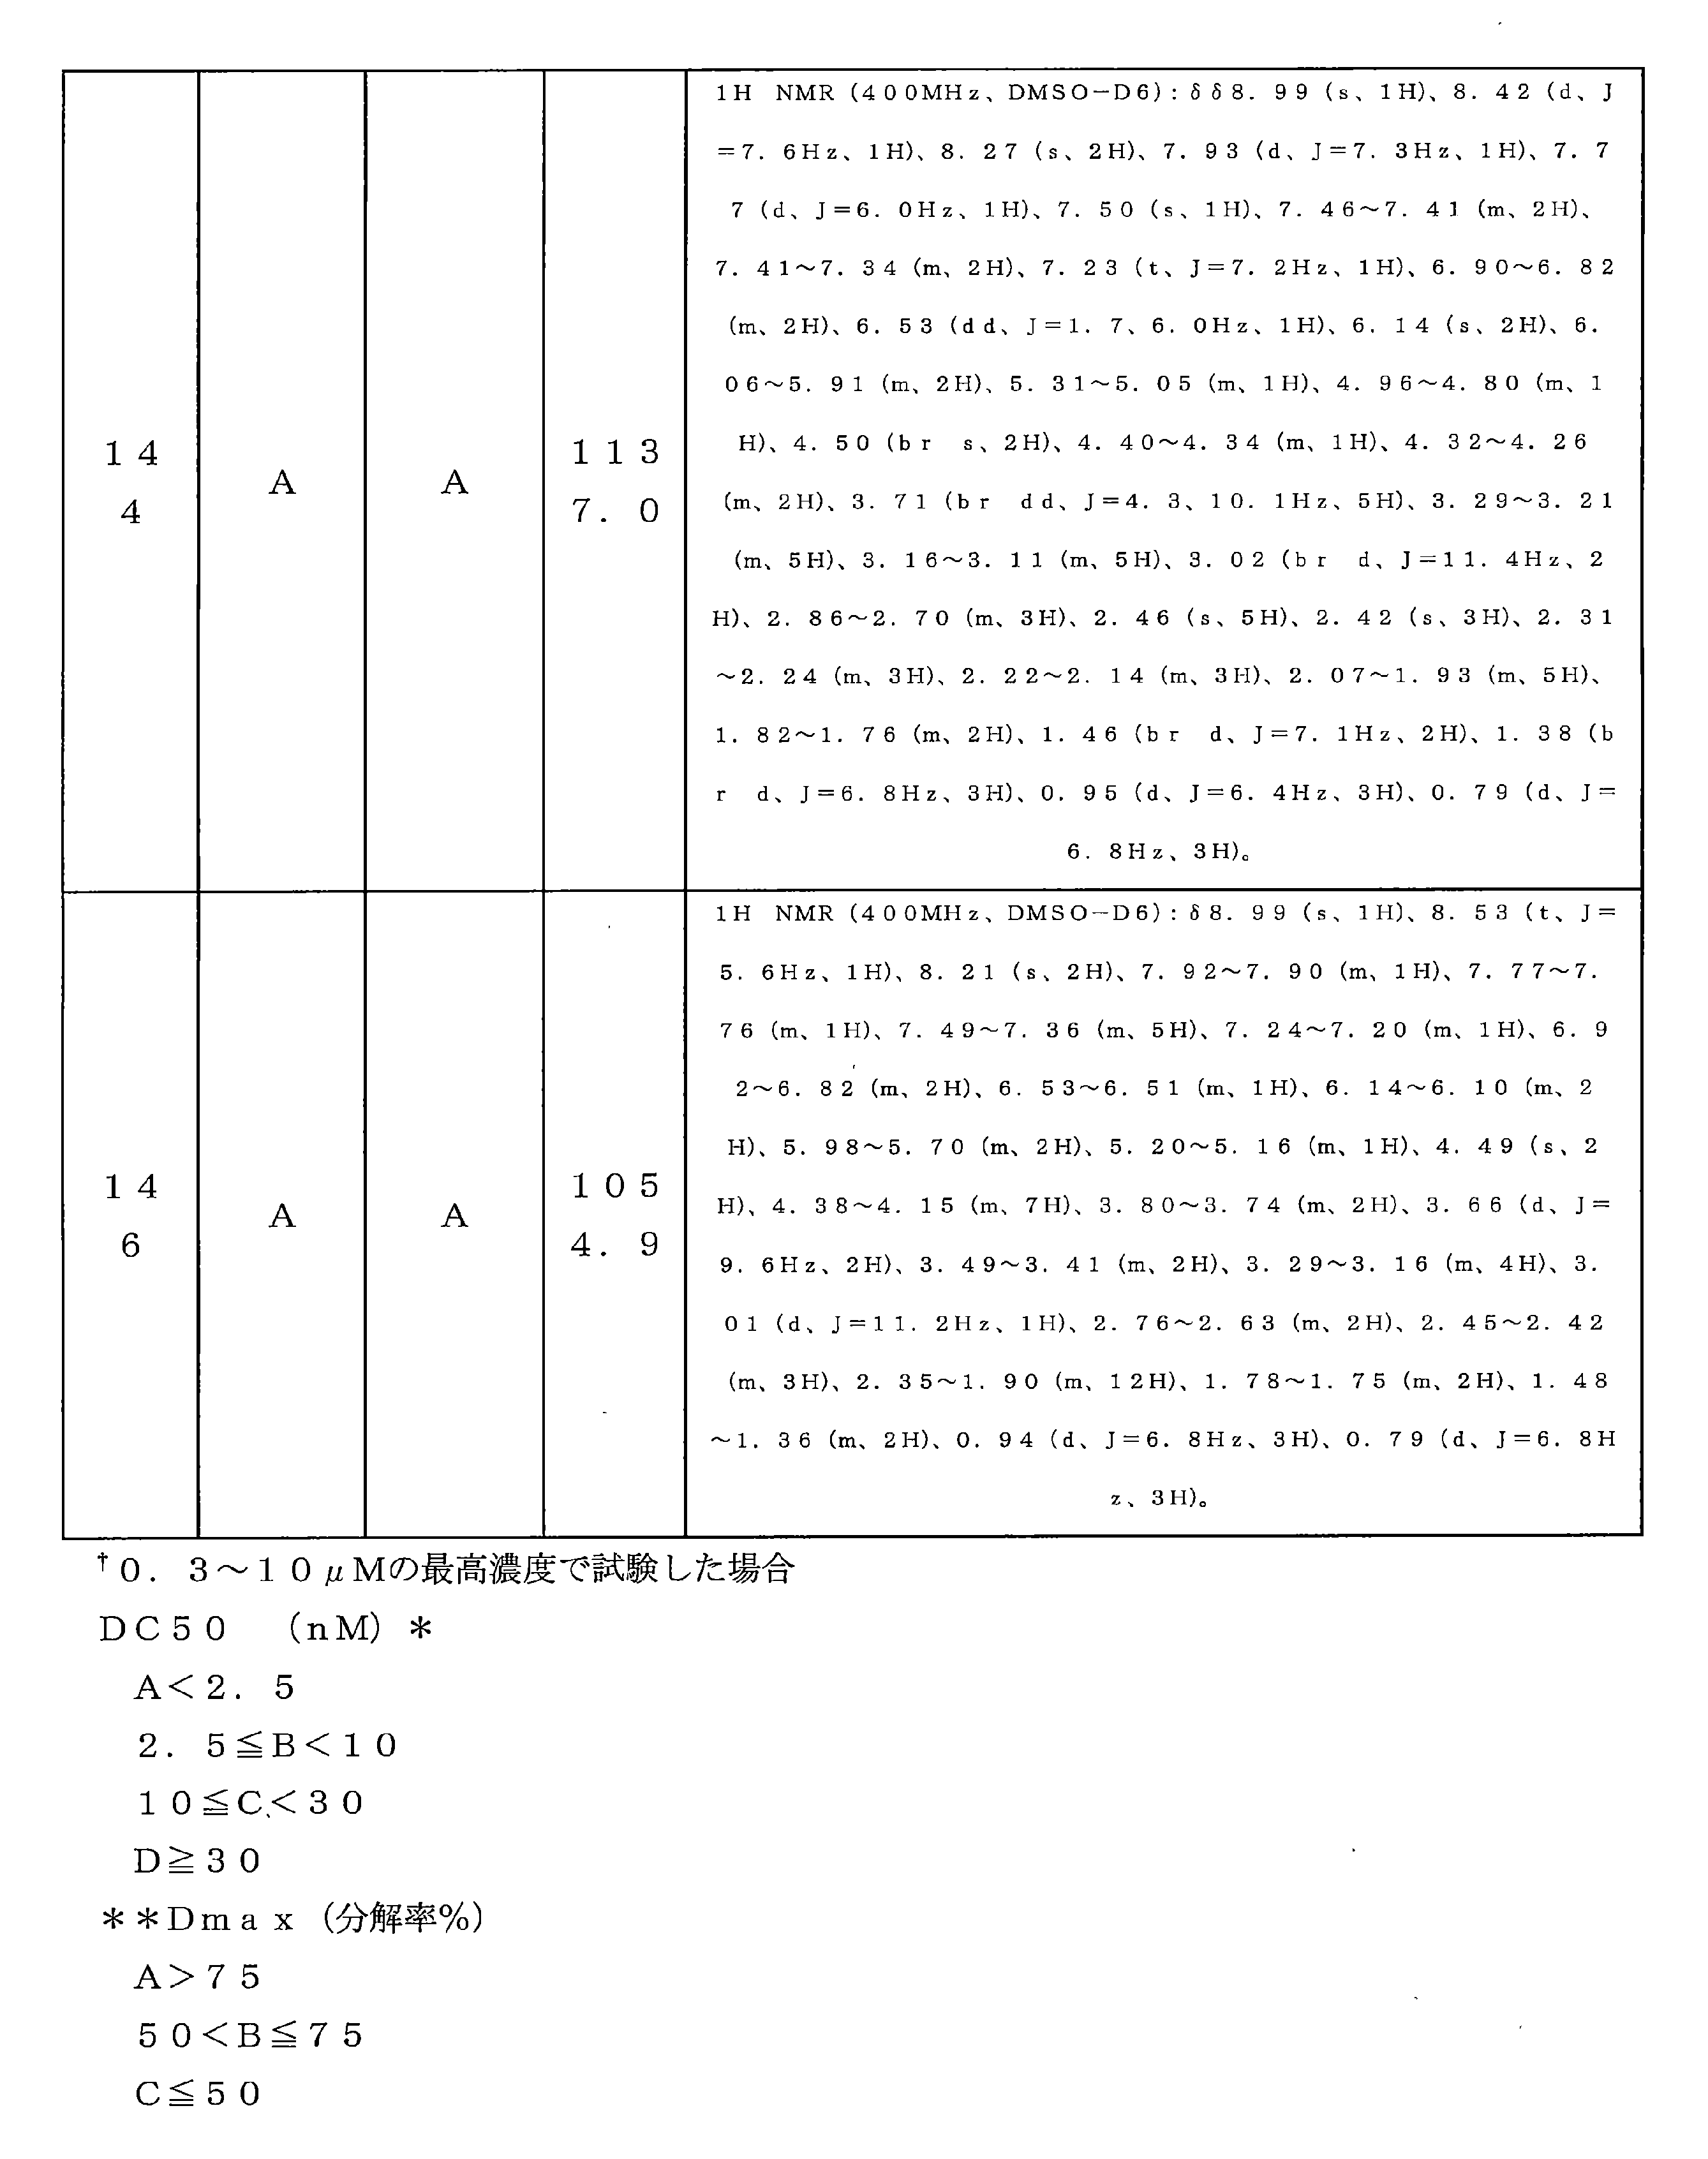
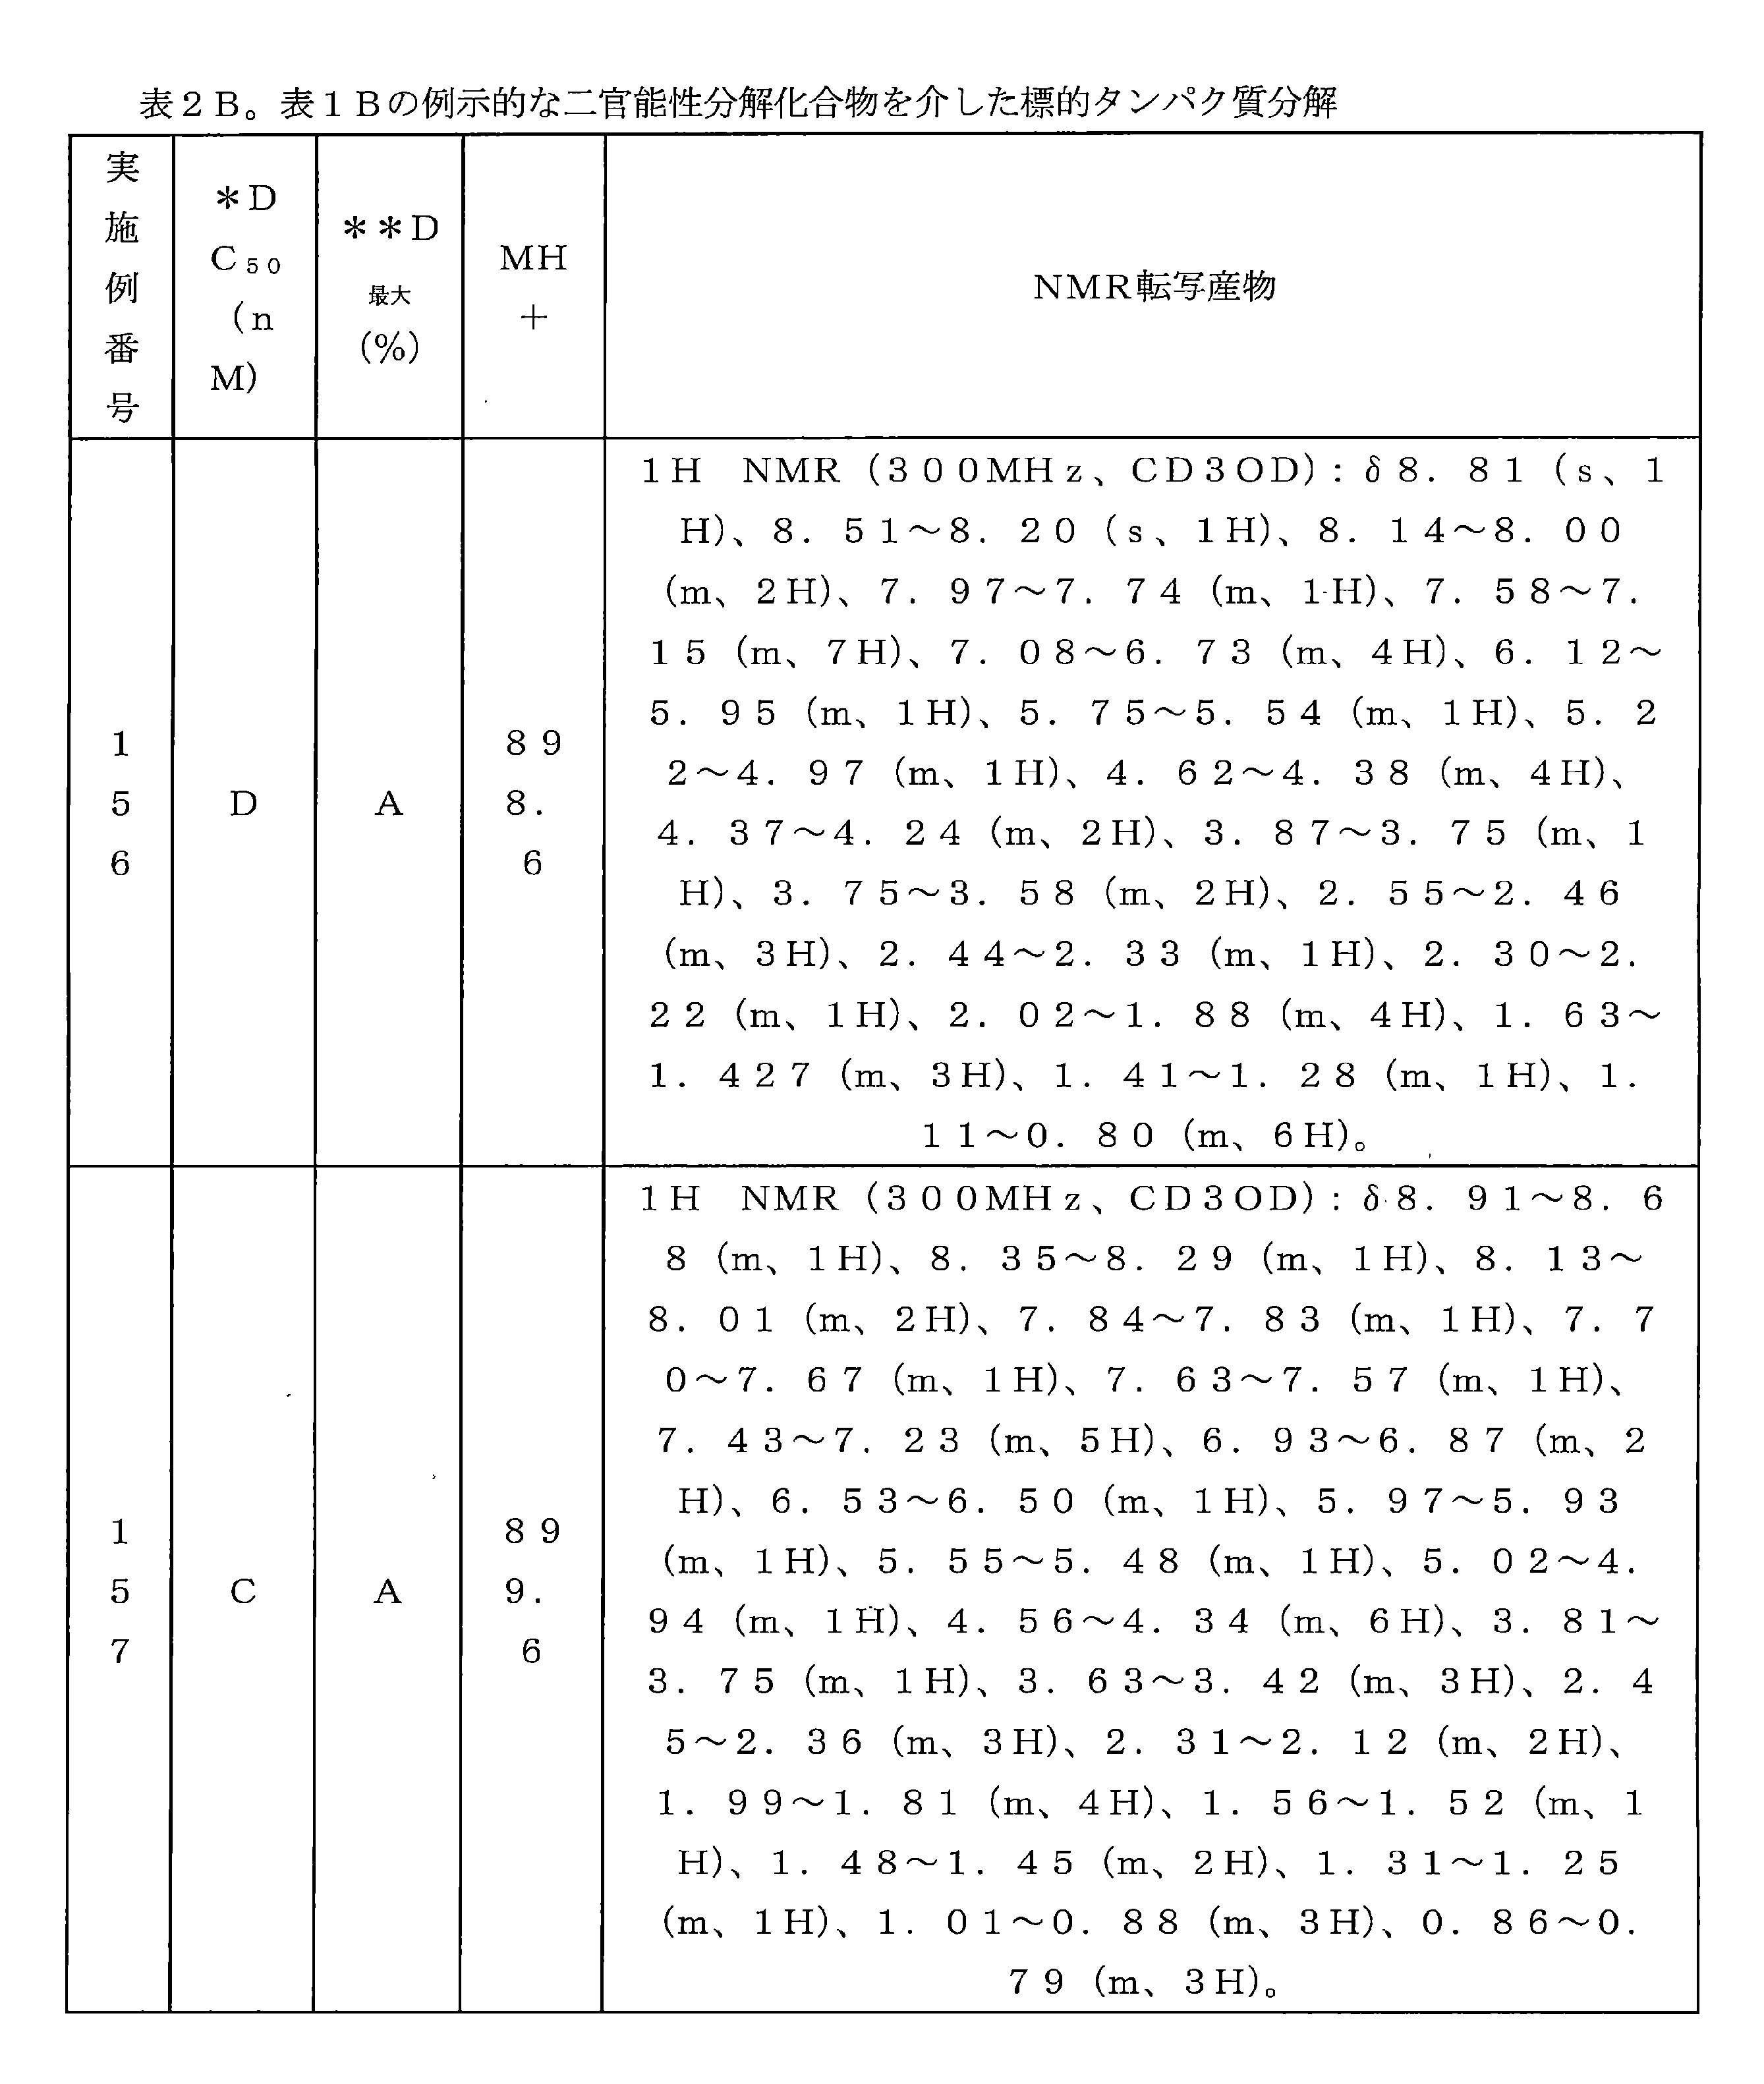


































SMARCA2、BRAHMA、またはBRMタンパク質動員部分およびE3リガーゼ動員部分を含む新規の二官能性分子が、PROTAC技術を通じて記載されている。本開示の二官能性分子は、SMARCA2を積極的に分解し、ロバストな細胞増殖抑制およびアポトーシス誘導をもたらす。PROTAC媒介タンパク質分解は、従来のアプローチによって病理学的タンパク質を「薬剤化不可能」に標的化する上で有望な戦略を提供する。 Novel bifunctional molecules containing SMARCA2, BRAHMA, or BRM protein recruitment moieties and E3 ligase recruitment moieties are described through PROTAC technology. The bifunctional molecules of the present disclosure actively degrade SMARCA2, resulting in robust cell growth inhibition and apoptosis induction. PROTAC-mediated protein degradation offers a promising strategy in targeting pathological proteins that are "undruggable" by conventional approaches.
本出願全体を通して引用されているすべての参考文献、特許、係属中の特許出願および公開された特許の内容は、参照により本明細書に明示的に組み込まれる。当業者は、本明細書に記述される本開示の特定の実施形態に対する多くの均等物を認識するか、または日常的な実験のみを使用して確認することができるであろう。かかる均等物は、以下の請求項に包含されることが意図される。本明細書に記載される詳細な実施例および実施形態は、解説目的のための例示としてのみ提供され、決して本開示を限定するものとはみなされないことを理解されたい。その観点での様々な改変または変更は、当分野の当業者に提案されるものであり、本出願の主旨および範囲内に含まれ、添付の請求項の範囲内で考慮される。例えば、成分の相対量は、望ましい効果を最適化するために変更されてもよく、追加的な成分が加えられてもよく、および/または類似した成分が、記載される成分のうちの1つ以上と置き換えられてもよい。本開示のシステム、方法、およびプロセスに関連付けられた追加的な有益な特徴および機能は、添付の請求項から明らかであろう。さらに当分野の当業者は、日常的な実験手法を越えない手法を使用して、本明細書に記載される本開示の特定の実施形態に対する多くの均等を認識するか、または確認することができるであろう。かかる均等物は、以下の請求項に包含されることが意図される。 The contents of all references, patents, pending patent applications and published patents cited throughout this application are expressly incorporated herein by reference. Those skilled in the art will recognize or be able to ascertain using no more than routine experimentation many equivalents to the specific embodiments of the present disclosure described herein. Such equivalents are intended to be encompassed by the following claims. It should be understood that the detailed examples and embodiments described herein are provided as examples only for illustrative purposes and are not to be considered as limiting the present disclosure in any way. Various modifications or alterations in that respect will be suggested to those skilled in the art and are within the spirit and scope of the present application and are contemplated within the scope of the appended claims. For example, the relative amounts of the components may be altered to optimize the desired effect, additional components may be added, and/or similar components may be substituted for one or more of the components described. Additional beneficial features and functions associated with the systems, methods, and processes of the present disclosure will be apparent from the appended claims. Moreover, those skilled in the art will recognize or be able to ascertain many equivalents to the specific embodiments of the present disclosure described herein using no more than routine experimentation. Such equivalents are intended to be encompassed by the following claims.
Claims (12)
PTM-L-ULM
を有する化合物、またはその薬学的に許容される塩であって、
式中、
前記ULMが、
式中、
R1が、C1-3アルキルであり;
R14aが、C1-3アルキルであり;
R14bが、HまたはC1-3アルキルであり;
R15が、
R28が、C1-3アルキルであり;
R3が、イソキサゾリルであり;
oが、0、1または2であり;
R16が、H、C1-4アルキル、フルオロ、クロロ、CN、またはC1-4アルコキシであり;
X4、X5、およびX6の各々が、CHであり;
前記Lが、
-(AL)q-
により表される前記ULMと前記PTMを連結する化学結合部分であり、
式中、
qが、1、2、3、4または5であり;
各ALが、独立して、CH2、O、シクロブチル、ピペリジニルまたはピペラジニルであり;
前記PTMが、
式中、
WPTM1が、ヒドロキシで任意に置換されたフェニルであり、
WPTM2が、アミノで置換されたピリダジニルであり、
WPTM3が、ピラゾリルまたは3,8-ジアザビシクロ[3.2.1]オクタニルであり、
WPTM4が、ピペリジニルであり、
LPTMが、C1-C3アルキレンであり、
PTM-L-ULM
or a pharma- ceutically acceptable salt thereof,
In the formula,
The ULM is
In the formula,
R 1 is C 1-3 alkyl;
R 14a is C 1-3 alkyl;
R 14b is H or C 1-3 alkyl;
R15 is
R 28 is C 1-3 alkyl;
R3 is isoxazolyl;
o is 0, 1 or 2;
R 16 is H, C 1-4 alkyl, fluoro, chloro, CN, or C 1-4 alkoxy;
Each of X 4 , X 5 , and X 6 is CH;
The L is
-( AL ) q-
is a chemical linking moiety linking the ULM and the PTM, represented by
In the formula,
q is 1, 2, 3, 4 or 5;
each A L is independently CH 2 , O, cyclobutyl, piperidinyl, or piperazinyl;
The PTM is
In the formula,
W PTM1 is phenyl optionally substituted with hydroxy;
W PTM2 is pyridazinyl substituted with amino;
W PTM3 is pyrazolyl or 3,8-diazabicyclo[3.2.1]octanyl;
W PTM4 is piperidinyl;
L PTM is C 1 -C 3 alkylene;
式中、
R30が、Hであり、
R28が、メチルである、請求項1に記載の化合物。 The ULM is
In the formula,
R 30 is H;
The compound of claim 1 , wherein R 28 is methyl.
Priority Applications (1)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| JP2024019396A JP2024056848A (en) | 2019-10-01 | 2024-02-13 | BRM TARGETING COMPOUNDS AND RELATED METHODS OF USE |
Applications Claiming Priority (3)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| US16/590,329 | 2019-10-01 | ||
| US16/590,329 US12539292B2 (en) | 2018-04-01 | 2019-10-01 | BRM targeting compounds and associated methods of use |
| PCT/US2020/053815 WO2021067606A1 (en) | 2019-10-01 | 2020-10-01 | Brm targeting compounds and associated methods of use |
Related Child Applications (1)
| Application Number | Title | Priority Date | Filing Date |
|---|---|---|---|
| JP2024019396A Division JP2024056848A (en) | 2019-10-01 | 2024-02-13 | BRM TARGETING COMPOUNDS AND RELATED METHODS OF USE |
Publications (2)
| Publication Number | Publication Date |
|---|---|
| JP2022550437A JP2022550437A (en) | 2022-12-01 |
| JP7502425B2 true JP7502425B2 (en) | 2024-06-18 |
Family
ID=72964828
Family Applications (2)
| Application Number | Title | Priority Date | Filing Date |
|---|---|---|---|
| JP2022520255A Active JP7502425B2 (en) | 2019-10-01 | 2020-10-01 | BRM TARGETING COMPOUNDS AND RELATED METHODS OF USE |
| JP2024019396A Pending JP2024056848A (en) | 2019-10-01 | 2024-02-13 | BRM TARGETING COMPOUNDS AND RELATED METHODS OF USE |
Family Applications After (1)
| Application Number | Title | Priority Date | Filing Date |
|---|---|---|---|
| JP2024019396A Pending JP2024056848A (en) | 2019-10-01 | 2024-02-13 | BRM TARGETING COMPOUNDS AND RELATED METHODS OF USE |
Country Status (4)
| Country | Link |
|---|---|
| EP (1) | EP4038066A1 (en) |
| JP (2) | JP7502425B2 (en) |
| CN (1) | CN114787159B (en) |
| WO (1) | WO2021067606A1 (en) |
Cited By (1)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| JP2024056848A (en) * | 2019-10-01 | 2024-04-23 | アルヴィナス・オペレーションズ・インコーポレイテッド | BRM TARGETING COMPOUNDS AND RELATED METHODS OF USE |
Families Citing this family (18)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| EP3737666A4 (en) | 2018-01-12 | 2022-01-05 | Kymera Therapeutics, Inc. | Protein degraders and uses thereof |
| EP3817822A4 (en) | 2018-07-06 | 2022-07-27 | Kymera Therapeutics, Inc. | PROTEIN DEGRADING AGENTS AND THEIR USES |
| WO2020251971A1 (en) | 2019-06-10 | 2020-12-17 | Kymera Therapeutics, Inc. | Smarca degraders and uses thereof |
| WO2020251972A1 (en) | 2019-06-10 | 2020-12-17 | Kymera Therapeutics, Inc. | Smarca degraders and uses thereof |
| US12569485B2 (en) | 2019-12-23 | 2026-03-10 | Kymera Therapeutics, Inc. | SMARCA inhibitors and uses thereof |
| CN115297931A (en) | 2019-12-23 | 2022-11-04 | 凯麦拉医疗公司 | SMARCA degrading agents and uses thereof |
| US12528814B2 (en) | 2020-02-19 | 2026-01-20 | Nurix Therapeutics, Inc. | Bifunctional degraders of interleukin-1 receptor-associated kinases and therapeutic use thereof |
| TW202216215A (en) * | 2020-07-21 | 2022-05-01 | 美商建南德克公司 | Antibody-conjugated chemical inducers of degradation of brm and methods thereof |
| EP4259144A4 (en) | 2020-12-09 | 2025-08-20 | Kymera Therapeutics Inc | SMARCA DECORATORS AND USES THEREOF |
| JP2024504932A (en) | 2021-01-13 | 2024-02-02 | モンテ ローザ セラピューティクス, インコーポレイテッド | isoindolinone compound |
| WO2023287787A1 (en) * | 2021-07-13 | 2023-01-19 | Prelude Therapeutics, Incorporated | Brm targeting compounds and associated methods of use |
| CA3225467A1 (en) * | 2021-08-09 | 2023-02-16 | Genentech, Inc. | Phenol derivatives for use in the modulation of brm |
| EP4367118B1 (en) | 2021-08-18 | 2025-01-29 | Nurix Therapeutics, Inc. | Bifunctional degraders of interleukin-1 receptor-associated kinases and therapeutic use thereof |
| CN118696045A (en) * | 2021-11-23 | 2024-09-24 | 南京再明医药有限公司 | BRM selective degradation agent compounds and their applications |
| CN118679160A (en) * | 2021-11-24 | 2024-09-20 | 阿维纳斯运营公司 | BRM targeting compounds and related methods of use |
| AR128330A1 (en) | 2022-01-26 | 2024-04-17 | Genentech Inc | CHEMICAL DEGRADATION INDUCERS CONJUGATED WITH ANTIBODY AND METHODS OF THESE |
| AR128331A1 (en) * | 2022-01-26 | 2024-04-17 | Genentech Inc | CHEMICAL DEGRADATION INDUCTORS CONJUGATED WITH ANTIBODIES AND METHODS OF THESE |
| WO2023152666A1 (en) * | 2022-02-09 | 2023-08-17 | Aurigene Oncology Limited | 3-substituted pyridazine compounds as smarca2 and/or smarca4 degraders |
Citations (2)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| JP2018510851A (en) | 2015-02-25 | 2018-04-19 | ジェネンテック, インコーポレイテッド | Therapeutic pyridazine compounds and uses thereof |
| JP2018512449A (en) | 2015-03-18 | 2018-05-17 | アルビナス インコーポレイテッド | Compounds and methods for improved degradation of target proteins |
Family Cites Families (9)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| US4522811A (en) | 1982-07-08 | 1985-06-11 | Syntex (U.S.A.) Inc. | Serial injection of muramyldipeptides and liposomes enhances the anti-infective activity of muramyldipeptides |
| AU2011338615B2 (en) | 2010-12-07 | 2017-07-27 | Yale University | Small-molecule hydrophobic tagging of fusion proteins and induced degradation of same |
| JP2015508414A (en) | 2012-01-12 | 2015-03-19 | イエール ユニバーシティ | Compounds and methods for enhanced degradation of target proteins and other polypeptides by E3 ubiquitin ligase |
| CN119591582A (en) | 2014-04-14 | 2025-03-11 | 阿尔维纳斯运营股份有限公司 | Imide-based proteolytic modulators and related methods of use |
| EP3337476A4 (en) | 2015-08-19 | 2019-09-04 | Arvinas, Inc. | COMPOUNDS AND METHODS FOR THE TARGETED REMOVAL OF BROMO-DOMAIN-CONTAINING PROTEINS |
| CN109152933B (en) * | 2016-04-21 | 2022-12-02 | 生物风险投资有限责任公司 | Compounds that induce degradation of anti-apoptotic BCL-2 family proteins and uses thereof |
| EP3577109A4 (en) | 2017-01-31 | 2020-11-18 | Arvinas Operations, Inc. | CEREBLON LIGANDS AND BIFUNCTIONAL CONNECTIONS THEREOF |
| KR20240130153A (en) * | 2018-04-01 | 2024-08-28 | 아비나스 오퍼레이션스, 인코포레이티드 | Brm targeting compounds and associated methods of use |
| WO2021067606A1 (en) * | 2019-10-01 | 2021-04-08 | Arvinas Operations, Inc. | Brm targeting compounds and associated methods of use |
-
2020
- 2020-10-01 WO PCT/US2020/053815 patent/WO2021067606A1/en not_active Ceased
- 2020-10-01 JP JP2022520255A patent/JP7502425B2/en active Active
- 2020-10-01 EP EP20793893.7A patent/EP4038066A1/en active Pending
- 2020-10-01 CN CN202080083758.XA patent/CN114787159B/en active Active
-
2024
- 2024-02-13 JP JP2024019396A patent/JP2024056848A/en active Pending
Patent Citations (2)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| JP2018510851A (en) | 2015-02-25 | 2018-04-19 | ジェネンテック, インコーポレイテッド | Therapeutic pyridazine compounds and uses thereof |
| JP2018512449A (en) | 2015-03-18 | 2018-05-17 | アルビナス インコーポレイテッド | Compounds and methods for improved degradation of target proteins |
Non-Patent Citations (1)
| Title |
|---|
| NATURE CHEMICAL BIOLOGY,2019年07月,Vol.15,p.672-680 |
Cited By (1)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| JP2024056848A (en) * | 2019-10-01 | 2024-04-23 | アルヴィナス・オペレーションズ・インコーポレイテッド | BRM TARGETING COMPOUNDS AND RELATED METHODS OF USE |
Also Published As
| Publication number | Publication date |
|---|---|
| CN114787159B (en) | 2024-09-27 |
| CN114787159A (en) | 2022-07-22 |
| JP2024056848A (en) | 2024-04-23 |
| WO2021067606A1 (en) | 2021-04-08 |
| EP4038066A1 (en) | 2022-08-10 |
| JP2022550437A (en) | 2022-12-01 |
Similar Documents
| Publication | Publication Date | Title |
|---|---|---|
| JP2024056848A (en) | BRM TARGETING COMPOUNDS AND RELATED METHODS OF USE | |
| US20240076295A1 (en) | Brm targeting compounds and associated methods of use | |
| US20260041671A1 (en) | Brm targeting compounds and associated methods of use | |
| JP7834774B2 (en) | Regulators of BCL6 proteolysis and related uses | |
| AU2019404934B2 (en) | Macrocyclic compounds and their use in the treatment of disease | |
| JP2024052757A (en) | Bifunctional molecules containing an E3 ubiquitin ligase binding moiety linked to a BCL6 targeting moiety | |
| US20230002397A1 (en) | Small molecule degraders of helios and metods of use | |
| ES2898069T3 (en) | N-(pyridin-2-yl)pyridinesulfonamide derivatives and their use in the treatment of disease | |
| IL284424B2 (en) | Tetrahydronaphthalene and tetrahydroisoquinoline derivatives as estrogen receptor degraders | |
| IL281188B1 (en) | Polycyclic compounds and methods for the targeted degradation of rapidly accelerated fibrosarcoma polypeptides | |
| IL283190B2 (en) | Heterocyclics that function as antibiotic agents | |
| EP2124951A1 (en) | 5-cyan0-4- (pyrrolo ý2, 3b¨pyridine-3-yl) -pyrimidine derivatives useful as protein kinase inhibitors | |
| US20210284624A1 (en) | Immunomodulatory compounds | |
| JP2025507762A (en) | Compounds and methods useful for stabilizing phenylalanine hydroxylase mutations - Patents.com | |
| TW202146384A (en) | Substituted aza five-membered ring compound and its application in medicine | |
| RU2797832C2 (en) | Compounds targeting brm and related uses thereof | |
| WO2024108009A1 (en) | Dyrk/clk protacs and uses thereof | |
| HK40095524A (en) | Pyrazoleamide derivatives | |
| HK40130773A (en) | Brm targeting compounds and associated methods of use | |
| HK40115036A (en) | Brm targeting compounds and associated methods of use | |
| HK40037668B (en) | Brm targeting compounds and associated methods of use | |
| TW201231467A (en) | Pyrazolopyridine kinase inhibitors |
Legal Events
| Date | Code | Title | Description |
|---|---|---|---|
| A621 | Written request for application examination |
Free format text: JAPANESE INTERMEDIATE CODE: A621 Effective date: 20220531 |
|
| A131 | Notification of reasons for refusal |
Free format text: JAPANESE INTERMEDIATE CODE: A131 Effective date: 20230619 |
|
| A521 | Request for written amendment filed |
Free format text: JAPANESE INTERMEDIATE CODE: A523 Effective date: 20230915 |
|
| A131 | Notification of reasons for refusal |
Free format text: JAPANESE INTERMEDIATE CODE: A131 Effective date: 20231113 |
|
| A521 | Request for written amendment filed |
Free format text: JAPANESE INTERMEDIATE CODE: A523 Effective date: 20240213 |
|
| TRDD | Decision of grant or rejection written | ||
| A01 | Written decision to grant a patent or to grant a registration (utility model) |
Free format text: JAPANESE INTERMEDIATE CODE: A01 Effective date: 20240513 |
|
| A61 | First payment of annual fees (during grant procedure) |
Free format text: JAPANESE INTERMEDIATE CODE: A61 Effective date: 20240606 |
|
| R150 | Certificate of patent or registration of utility model |
Ref document number: 7502425 Country of ref document: JP Free format text: JAPANESE INTERMEDIATE CODE: R150 |