WO2000035901A1 - Chromanderivate - Google Patents
Chromanderivate Download PDFInfo
- Publication number
- WO2000035901A1 WO2000035901A1 PCT/EP1999/009333 EP9909333W WO0035901A1 WO 2000035901 A1 WO2000035901 A1 WO 2000035901A1 EP 9909333 W EP9909333 W EP 9909333W WO 0035901 A1 WO0035901 A1 WO 0035901A1
- Authority
- WO
- WIPO (PCT)
- Prior art keywords
- formula
- chroman
- compounds
- ylmethyl
- acid
- Prior art date
- Legal status (The legal status is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the status listed.)
- Ceased
Links
- 0 CC1(*)/C=C(/C(C=C(CN(*)*)O2)=*)\C2=C/C(C)(*)/C=C1 Chemical compound CC1(*)/C=C(/C(C=C(CN(*)*)O2)=*)\C2=C/C(C)(*)/C=C1 0.000 description 1
- SJFFVMCXWBBLGN-WDIFMUJUSA-N CC[C@@H](CCC1)P1c(cccc1)c1P1[C@@H](CC)CC[C@@H]1CC Chemical compound CC[C@@H](CCC1)P1c(cccc1)c1P1[C@@H](CC)CC[C@@H]1CC SJFFVMCXWBBLGN-WDIFMUJUSA-N 0.000 description 1
- RIULASLQEIKCJM-PELVSDQASA-N C[C@@H](C12)C11C=CC1C[C@@H]2P(c1ccccc1)c1ccccc1 Chemical compound C[C@@H](C12)C11C=CC1C[C@@H]2P(c1ccccc1)c1ccccc1 RIULASLQEIKCJM-PELVSDQASA-N 0.000 description 1
Classifications
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D405/00—Heterocyclic compounds containing both one or more hetero rings having oxygen atoms as the only ring hetero atoms, and one or more rings having nitrogen as the only ring hetero atom
- C07D405/02—Heterocyclic compounds containing both one or more hetero rings having oxygen atoms as the only ring hetero atoms, and one or more rings having nitrogen as the only ring hetero atom containing two hetero rings
- C07D405/12—Heterocyclic compounds containing both one or more hetero rings having oxygen atoms as the only ring hetero atoms, and one or more rings having nitrogen as the only ring hetero atom containing two hetero rings linked by a chain containing hetero atoms as chain links
-
- A—HUMAN NECESSITIES
- A61—MEDICAL OR VETERINARY SCIENCE; HYGIENE
- A61P—SPECIFIC THERAPEUTIC ACTIVITY OF CHEMICAL COMPOUNDS OR MEDICINAL PREPARATIONS
- A61P25/00—Drugs for disorders of the nervous system
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D311/00—Heterocyclic compounds containing six-membered rings having one oxygen atom as the only hetero atom, condensed with other rings
- C07D311/02—Heterocyclic compounds containing six-membered rings having one oxygen atom as the only hetero atom, condensed with other rings ortho- or peri-condensed with carbocyclic rings or ring systems
- C07D311/04—Benzo[b]pyrans, not hydrogenated in the carbocyclic ring
- C07D311/22—Benzo[b]pyrans, not hydrogenated in the carbocyclic ring with oxygen or sulfur atoms directly attached in position 4
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D311/00—Heterocyclic compounds containing six-membered rings having one oxygen atom as the only hetero atom, condensed with other rings
- C07D311/02—Heterocyclic compounds containing six-membered rings having one oxygen atom as the only hetero atom, condensed with other rings ortho- or peri-condensed with carbocyclic rings or ring systems
- C07D311/04—Benzo[b]pyrans, not hydrogenated in the carbocyclic ring
- C07D311/58—Benzo[b]pyrans, not hydrogenated in the carbocyclic ring other than with oxygen or sulphur atoms in position 2 or 4
-
- Y—GENERAL TAGGING OF NEW TECHNOLOGICAL DEVELOPMENTS; GENERAL TAGGING OF CROSS-SECTIONAL TECHNOLOGIES SPANNING OVER SEVERAL SECTIONS OF THE IPC; TECHNICAL SUBJECTS COVERED BY FORMER USPC CROSS-REFERENCE ART COLLECTIONS [XRACs] AND DIGESTS
- Y02—TECHNOLOGIES OR APPLICATIONS FOR MITIGATION OR ADAPTATION AGAINST CLIMATE CHANGE
- Y02P—CLIMATE CHANGE MITIGATION TECHNOLOGIES IN THE PRODUCTION OR PROCESSING OF GOODS
- Y02P20/00—Technologies relating to chemical industry
- Y02P20/50—Improvements relating to the production of bulk chemicals
- Y02P20/55—Design of synthesis routes, e.g. reducing the use of auxiliary or protecting groups
Definitions
- the invention relates to chroman derivatives of the formula I.
- R 1 acyl with 1-6 C atoms, -CO-R 5 or an amino protecting group
- R 2 H or alkyl with 1-6 C atoms
- R 3 , R 4 each independently of one another H, alkyl having 1-6 C atoms, CN,
- the invention also relates to the optically active forms, the racemates, the enantiomers and the hydrates and solvates, e.g. Alcoholates, these compounds.
- the invention was based on the object of finding new compounds which can be used in particular as intermediates in the synthesis of medicaments.
- the invention relates to the chroman derivatives of the formula I and their salts
- alkyl has 1 to 6, preferably 1, 2, 3 or
- alkyl preferably means methyl or ethyl, furthermore propyl, isopropyl, and also butyl isobutyl sec-butyl or tert-butyl acyl has 1 to 6, preferably 1, 2, 3 or 4 carbon atoms, acyl means in particular acetyl, propionyl or butyryl?
- R is preferably H, and also methyl, ethyl or propyl
- R 3 and R 4 are preferably H.
- R 5 is preferably, for example, phenyl, o-, m- or p-tolyl, o-, m- or p-
- the radical R 1 means acyl, -CO-R 5 or a known one
- amino protecting group is generally known and refers to groups which are suitable for protecting (blocking) an amino group from chemical reactions, but which are easily removable after the desired chemical reaction has been carried out elsewhere in the molecule unsubstituted acyl, aryl, aralkoxymethyl or aralkyl groups are particularly suitable for such groups.
- acyl group is to be understood in the broadest sense in connection with the present process and the present compounds or heterocydic carboxylic acids or sulfonic acids derived acyl groups and especially alkoxycarbonyl, aryloxycarbonyl and especially aralkoxycarbonyl groups.
- alkanoyl such as acetyl, propionyl, butyryl
- Aralkanoyi such as phenylacetyl
- ⁇ such as benzoyl or toluyl; Aryloxyalkanoyl such as phenoxyacetyl; Alkoxycarbonyl such as methoxycarbonyl, ethoxycarbonyl, 2,2,2-trichloroethoxycarbonyl, BOC (tert-butoxycarbonyl), 2-iodoethoxycarbonyl; Aralkyloxycarbonyl such as CBZ (carbobenzoxycarbonyi, also called "Z”), 4-methoxy
- benzyloxycarbonyl, FMOC (9-fluorenylmethoxycarbonyl); Arylsulfonyl such as Mtr (4-methoxy-2,3,6-trimethylphenylsulfonyl).
- Preferred amino protective groups are BOC and Mtr, furthermore CBZ or FMOC.
- the compounds of formula I can have one or more chiral centers
- the invention further relates to a process for the preparation of 20 chroman derivatives of the formula I according to Claim 1 and salts thereof, in which X is O, characterized in that a compound of the formula II
- the invention also relates to a process for the preparation of chroman derivatives of the formula I as claimed in claim 1 and salts thereof, where X is H, 35901
- R 1 , R 2 , R 3 , R 4 have the meanings given in claim 1 and X is 0, hydrogenated with the aid of an enantiomerically enriched catalyst, and then reduced as usual.
- (2-acetylaminomethyl) -chromen-4-one can be hydrogenated with various enantiomerically pure rhodium-diphosphine complexes to give enantiomerically enriched (2-acetylaminomethyl) -chroman-4-one.
- the invention also relates to a process for the preparation of chroman derivatives of the formula I, characterized in that the enantiomer-enriched catalyst is a transition metal complex.
- the catalyst is particularly preferably a transition metal complex containing a metal selected from the group consisting of rhodium, iridium, ruthenium and palladium.
- the invention further relates to a process for the preparation of chroman derivatives of the formula I, characterized in that the catalyst is a transition metal complex in which the transition metal is complexed with a chiral diphosphine ligand.
- the (R) - or (S) -enantiomer of the ligand in the catalyst is obtained in excess.
- Rh (COD) 2 OTf rhodium-cyclooctadiene triflate
- [Rh (COD) CI] 2 Rh (COD) 2 BF 4
- [lr (COD) CI] 2 serve as precursors for the chiral ligands, lr (COD) 2 BF 4 or [Ru (COD) CI 2 ] x .
- the compounds of the formula I and also the starting materials for their preparation are otherwise prepared by methods known per se, as described in the literature (for example in the standard works such as Houben-Weyl, methods of organic chemistry, Georg-Thieme-Verlag, Stuttgart) are described, namely under reaction conditions which are known and suitable for the reactions mentioned. Use can also be made of variants which are known per se and are not mentioned here in detail.
- the starting materials can also be formed in situ, so that they are not isolated from the reaction mixture, but instead are immediately reacted further to give the compounds of the formula I.
- Suitable inert solvents are, for example, Hydrocarbons like
- Sulfoxides such as dimethyl sulfoxide (DMSO); Carbon disulfide; Nitrover bonds such as nitromethane or nitrobenzene; Esters such as ethyl acetate, optionally also mixtures of the solvents mentioned with one another or mixtures with water.
- DMSO dimethyl sulfoxide
- Carbon disulfide Nitrover bonds such as nitromethane or nitrobenzene
- Esters such as ethyl acetate, optionally also mixtures of the solvents mentioned with one another or mixtures with water.
- the reaction time of the enantioselective hydrogenation is between a few minutes and 14 days, the reaction temperature is between 0 and 150 °, normally between 20 and 130 °.
- the catalyst / substrate ratio is usually between 1: 2000 and 1:50, particularly preferably between 1: 1000 and 1: 100.
- the response time is then e.g. between 3 and 20 hours.
- the hydrogenation is carried out under 1-200 bar of hydrogen, preferably at 3-100 bar.
- H is carried out according to the invention with hydrogen gas with the aid of an enantiomerically enriched catalyst in an inert solvent such as methanol or ethanol as described above, followed by a conversion of the 4-oxo group into a methylene group according to known conditions.
- the reduction is preferably carried out with hydrogen gas with transition metal catalysis (for example by hydrogenation on Raney nickel or Pd carbon in an inert
- Solvents such as methanol or ethanol).
- a radical R is split off from a compound of the formula I, for example using strong acids, expediently using TFA (trifluoroacetic acid) or perchloric acid, but also using other strong inorganic acids such as hydrochloric acid or sulfuric acid.
- strong acids expediently using TFA (trifluoroacetic acid) or perchloric acid, but also using other strong inorganic acids such as hydrochloric acid or sulfuric acid.
- strong organic carboxylic acids such as trichloroacetic acid or sulfonic acids such as benzene or p-toluenesulfonic acid.
- the presence of an additional inert solvent is possible, but not always necessary.
- Suitable inert solvents are preferably organic, for example carboxylic acids such as acetic acid, ethers such as tetrahydrofuran or dioxane, amides such as dimethylformamide, halogenated hydrocarbons such as dichloromethane, and also alcohols such as methanol, ethanol or isopropanol and water. Mixtures of the abovementioned solvents are also suitable. TFA is preferably used in excess without the addition of another solvent, perchloric acid in the form of a mixture of acetic acid and 70% perchloric acid in a ratio of 9: 1.
- the reaction temperatures are advantageously between about 0 and about 50 °, preferably between 15 and 30 ° .
- the BOC group is preferably split off with TFA in dichloromethane or with about 3 to 5 N hydrochloric acid in dioxane at 5-30 °.
- the acetyl group is split off by customary methods (P.J. Kocienski, Protecting Groups, Georg Thieme Verlag, Stuttgart, 1994).
- Hydrogenolytically removable protective groups can e.g. by treatment with hydrogen in the presence of a catalyst (e.g. a noble metal catalyst such as palladium, suitably on a support such as coal).
- a catalyst e.g. a noble metal catalyst such as palladium, suitably on a support such as coal.
- Suitable solvents are those given above, in particular e.g. Alcohols like methanol or ethanol or amides like DMF.
- the hydrogenolysis is generally carried out at temperatures between about 0 and 100 ° and pressures between about 1 and 200 bar, preferably at 20-30 ° and 1-10 bar.
- a base of the formula I can be converted into the associated acid addition salt using an acid, for example by reacting equivalent amounts of the base and the acid in an inert solvent such as ethanol and subsequent evaporation.
- acids that provide physiologically acceptable salts are suitable for this implementation.
- Inorganic acids can be used, for example sulfuric acid, nitric acid, hydrohalic acids such as hydrochloric acid or hydrobromic acid, phosphoric acids such as
- Formic acid acetic acid, propionic acid, pivalic acid, diethylacetic acid, malonic acid, succinic acid, pimelic acid, fumaric acid, maleic acid, lactic acid, tartaric acid, malic acid, citric acid, gluconic acid, ascorbic acid, nicotinic acid, isonicotinic acid, methane or hydroxysulfonic acid, ethane-sulfonic acid, ethanesulfonic acid, ethanesulfonic acid, ethanesulfonic acid, ethanesulfonic acid, ethanesulfonic acid, ethanesulfonic acid, ethanesulfonic acid, ethanesulfonic acid, ethanesulfonic acid, ethanesulfonic acid, ethanesulfonic acid, ethanesulfonic acid, ethanesulfonic acid, ethanesulfonic acid
- compounds of formula I with bases can be converted into the corresponding metal, in particular alkali metal or alkaline earth metal, or into the corresponding ammonium salts.
- the invention furthermore relates to the use of the compounds of the formula I as intermediates for the synthesis of medicaments.
- Corresponding medicaments are described, for example, in EP 0 707 007.
- the invention relates in particular to
- R 1 has the meaning given in claim 1.
- R 2 , R 3 and R 4 are H and X is hydrogen, hydrogenated with the aid of an enantiomer-enriched catalyst, b) that from the resulting enantiomer-enriched mixture of the (R) and (S) compounds of the formula I, in which
- R 1 has the meaning given in claim 1,
- R 2 , R 3 and R 4 are H and XO is the enantiomerically pure (R) compound of the formula I in which
- R 1 has the meaning given in claim 1,
- R 2 , R 3 and R 4 are H and X is 0, obtained by crystallization that c) the enantiomerically pure (R) compound of the formula I, in which R 1 has the meaning given in claim 1,
- R 2 , R 3 and R 4 are H and X is 0, then reduced as usual, that d) from the (R) compound of the formula I thus obtained, in which R 1 has the meaning given in claim 1,
- R 2 , R 3 and R 4 are H and XH, H is, the radical R 1 is split off that e) the (R) - (chroman-2-ylmethyl) amine thus obtained is converted into its acid addition salt and this is converted in a known manner to (R) -2- [5- (4-fluorophenyl) -3-pyridylmethylaminomethyl] -chroman and optionally converted into its acid addition salt, it being possible for the (R) enantiomer to be obtained by crystallization from the enantiomerically enriched (R, S) mixture also after stage c) or after stage d).
- the invention further relates to the use of the compounds of the formula I as intermediates for the synthesis of medicaments which have effects on the central nervous system.
- customary work-up means: if necessary, water is added, and if necessary, depending on the constitution of the end product, the pH is adjusted to between 2 and 10, extracted with ethyl acetate or dichloromethane, separated off, dries the organic phase over sodium sulfate, evaporates and purifies by chromatography on silica gel and / or by crystallization. Rf values on silica gel.
- Rh (COD) 2 OTf rhodium cyclooctadiene triflate
- the catalyst-substrate solution to be hydrogenated was introduced into an autoclave in a countercurrent flow of protective gas.
- the protective gas atmosphere was replaced by repeated flushing with hydrogen (1-5 bar H 2 atmosphere).
- the catalysts analog 1.2. gave the best results at 50 ° C and 80 bar hydrogen. As a rule, the hydrogenation was stopped after 15 hours.
- the keto group was reduced using palladium-carbon as a one-pot process.
- the crude ketone solution resulting from the homogeneous hydrogenation was admixed with 10% by weight of water-moist palladium-carbon (e.g. to 1 g of (2-acetylaminomethyl) -chrome ⁇ -4-one 100 mg of water-moist Pd / C) and 1ml of glacial acetic acid. At 7 bar hydrogen pressure and 50 ° C., hydrogenation was carried out for 14 hours.
- the palladium-carbon was removed by filtration.
- Elab complex metal anion Lsm pressure% ee no. Lig (additive)
Landscapes
- Chemical & Material Sciences (AREA)
- Organic Chemistry (AREA)
- Health & Medical Sciences (AREA)
- Nuclear Medicine, Radiotherapy & Molecular Imaging (AREA)
- Pharmacology & Pharmacy (AREA)
- Neurosurgery (AREA)
- Biomedical Technology (AREA)
- Chemical Kinetics & Catalysis (AREA)
- General Chemical & Material Sciences (AREA)
- Medicinal Chemistry (AREA)
- Bioinformatics & Cheminformatics (AREA)
- Engineering & Computer Science (AREA)
- Neurology (AREA)
- Life Sciences & Earth Sciences (AREA)
- Animal Behavior & Ethology (AREA)
- General Health & Medical Sciences (AREA)
- Public Health (AREA)
- Veterinary Medicine (AREA)
- Pharmaceuticals Containing Other Organic And Inorganic Compounds (AREA)
- Organic Low-Molecular-Weight Compounds And Preparation Thereof (AREA)
- Plural Heterocyclic Compounds (AREA)
- Pyrane Compounds (AREA)
- Low-Molecular Organic Synthesis Reactions Using Catalysts (AREA)
Abstract
Description


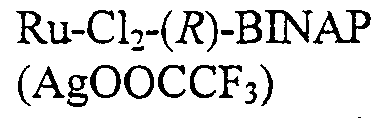
Claims


Priority Applications (16)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| UA2001075094A UA72235C2 (uk) | 1998-12-17 | 1999-01-12 | Похідні хроману |
| DE59904407T DE59904407D1 (en) | 1998-12-17 | 1999-12-01 | Chromanderivate |
| HK02104515.6A HK1043119B (zh) | 1998-12-17 | 1999-12-01 | 苯並二氫吡喃衍生物 |
| AU20946/00A AU760501B2 (en) | 1998-12-17 | 1999-12-01 | Chroman derivatives |
| AT99965422T ATE233253T1 (de) | 1998-12-17 | 1999-12-01 | Chromanderivate |
| SK816-2001A SK285837B6 (sk) | 1998-12-17 | 1999-12-01 | Deriváty chrómanu, spôsob ich prípravy a použitie |
| DK99965422T DK1140890T3 (da) | 1998-12-17 | 1999-12-01 | Chromanderivater. |
| US09/868,704 US6646136B1 (en) | 1998-12-17 | 1999-12-01 | Chroman derivatives |
| HU0104819A HU229060B1 (hu) | 1998-12-17 | 1999-12-01 | Krománszármazékok, azok alkalmazása és eljárás ilyen származékok elõállítására |
| SI9930269T SI1140890T1 (en) | 1998-12-17 | 1999-12-01 | Chroman derivatives |
| JP2000588161A JP4790910B2 (ja) | 1998-12-17 | 1999-12-01 | クロマン誘導体 |
| EP99965422A EP1140890B1 (de) | 1998-12-17 | 1999-12-01 | Chromanderivate |
| PL348191A PL194157B1 (pl) | 1998-12-17 | 1999-12-01 | Pochodne chromanu oraz sposób ich otrzymywania |
| BRPI9916188-5A BR9916188B1 (pt) | 1998-12-17 | 1999-12-01 | derivados de cromano. |
| CA002355098A CA2355098C (en) | 1998-12-17 | 1999-12-01 | Chroman derivatives |
| NO20012964A NO328132B1 (no) | 1998-12-17 | 2001-06-15 | Kromanderivater, fremgangsmater for fremstilling derav, anvendelse av forbindelsene som mellomprodukter for syntese av legemidler, samt fremgangsmate for fremstilling av et bestemt kromanderivat og dets salter. |
Applications Claiming Priority (2)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| DE19858341A DE19858341A1 (de) | 1998-12-17 | 1998-12-17 | Chromanderivate |
| DE19858341.9 | 1998-12-17 |
Publications (1)
| Publication Number | Publication Date |
|---|---|
| WO2000035901A1 true WO2000035901A1 (de) | 2000-06-22 |
Family
ID=7891486
Family Applications (1)
| Application Number | Title | Priority Date | Filing Date |
|---|---|---|---|
| PCT/EP1999/009333 Ceased WO2000035901A1 (de) | 1998-12-17 | 1999-12-01 | Chromanderivate |
Country Status (26)
| Country | Link |
|---|---|
| US (1) | US6646136B1 (de) |
| EP (1) | EP1140890B1 (de) |
| JP (1) | JP4790910B2 (de) |
| KR (1) | KR100676017B1 (de) |
| CN (1) | CN1140522C (de) |
| AR (1) | AR021900A1 (de) |
| AT (1) | ATE233253T1 (de) |
| AU (1) | AU760501B2 (de) |
| BR (1) | BR9916188B1 (de) |
| CA (1) | CA2355098C (de) |
| CZ (1) | CZ300544B6 (de) |
| DE (2) | DE19858341A1 (de) |
| DK (1) | DK1140890T3 (de) |
| ES (1) | ES2193779T3 (de) |
| HK (1) | HK1043119B (de) |
| HU (1) | HU229060B1 (de) |
| ID (1) | ID29273A (de) |
| NO (1) | NO328132B1 (de) |
| PL (1) | PL194157B1 (de) |
| PT (1) | PT1140890E (de) |
| RU (1) | RU2233277C2 (de) |
| SK (1) | SK285837B6 (de) |
| TW (1) | TWI222866B (de) |
| UA (1) | UA72235C2 (de) |
| WO (1) | WO2000035901A1 (de) |
| ZA (1) | ZA200105840B (de) |
Cited By (1)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| US6903227B2 (en) | 2000-06-02 | 2005-06-07 | Millennium Pharmaceuticals, Inc. | Synthesis of 2-acyl substituted chromanes and intermediates thereof |
Families Citing this family (11)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| DE10044091A1 (de) * | 2000-09-07 | 2002-04-04 | Merck Patent Gmbh | Chromanonderivate |
| DE60333094D1 (de) * | 2002-04-17 | 2010-08-05 | Cytokinetics Inc | Verbindungen, zusammensetzungen und verfahren |
| EP1539727B1 (de) * | 2002-07-17 | 2009-02-18 | Cytokinetics, Inc. | Verbindungen, zusammensetzungen und verfahren zur behandlung von zellulären proliferativen erkrankungen |
| JP2005539062A (ja) * | 2002-09-13 | 2005-12-22 | サイトキネティクス・インコーポレーテッド | 化合物、組成物および方法 |
| EP1670456A2 (de) * | 2003-10-06 | 2006-06-21 | Cytokinetics, Inc. | Verbindungen, zusammensetzungen und verfahren |
| CN100424034C (zh) * | 2006-08-25 | 2008-10-08 | 华中科技大学 | 一种磷石膏复合砖及其生产方法 |
| RU2404174C2 (ru) * | 2008-12-08 | 2010-11-20 | ГОУ ВПО "Северо-Осетинский государственный университет имени Коста Левановича Хетагурова" | 4-иминометил-3-(7-ацетоксихромон-3-ил) бензойная кислота, обладающая антибактериальной активностью |
| ES2534318B1 (es) * | 2013-10-18 | 2016-01-28 | Artax Biopharma Inc. | Derivados de cromeno sustituidos por alcóxido como inhibidores de la interacción TCR-Nck |
| ES2534336B1 (es) * | 2013-10-18 | 2016-01-28 | Artax Biopharma Inc. | Derivados de cromeno como inhibidores de la interacción TCR-Nck |
| CN108690017B (zh) * | 2018-05-04 | 2021-06-18 | 新乡学院 | 一种铑催化的莫西沙星侧链中间体的制备方法 |
| CN113979983B (zh) * | 2021-11-03 | 2023-10-03 | 中国人民解放军空军军医大学 | 一种非达司他关键中间体的不对称合成方法 |
Citations (2)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| EP0540914A1 (de) * | 1991-10-28 | 1993-05-12 | Bayer Ag | 2-Aminomethyl-chromane als Wirkstoffe in Arzneimitteln zur Behandlung von Erkrankungen des Zentralnervensystems |
| EP0707007A1 (de) * | 1994-10-14 | 1996-04-17 | MERCK PATENT GmbH | ZNS wirksame Amino(thio)ether-Derivate |
Family Cites Families (5)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| DE2745305A1 (de) * | 1977-10-07 | 1979-04-19 | Bayer Ag | Insektizide und akarizide mittel |
| DE2745306A1 (de) * | 1977-10-07 | 1979-04-19 | Bayer Ag | Insektizide mittel |
| CA2022693A1 (en) * | 1989-08-05 | 1991-02-06 | Rolf Gericke | Chroman derivatives |
| DE4102103A1 (de) * | 1991-01-25 | 1992-08-20 | Bayer Ag | Substituierte benzoxazepine und benzthiazepine, verfahren zu ihrer herstellung und ihre verwendung in arzneimitteln |
| RU2039739C1 (ru) * | 1993-03-01 | 1995-07-20 | Пятигорский фармацевтический институт | Замещенные фениламиды -3-(4`-оксо- 4н-1`-бензопиранил-3`)-пропен -2-карбоновой кислоты, обладающие антиаллергической активностью |
-
1998
- 1998-12-17 DE DE19858341A patent/DE19858341A1/de not_active Withdrawn
-
1999
- 1999-01-12 UA UA2001075094A patent/UA72235C2/uk unknown
- 1999-12-01 AT AT99965422T patent/ATE233253T1/de active
- 1999-12-01 ES ES99965422T patent/ES2193779T3/es not_active Expired - Lifetime
- 1999-12-01 AU AU20946/00A patent/AU760501B2/en not_active Expired
- 1999-12-01 CN CNB998142247A patent/CN1140522C/zh not_active Expired - Lifetime
- 1999-12-01 PL PL348191A patent/PL194157B1/pl unknown
- 1999-12-01 CA CA002355098A patent/CA2355098C/en not_active Expired - Fee Related
- 1999-12-01 KR KR1020017007512A patent/KR100676017B1/ko not_active Expired - Fee Related
- 1999-12-01 ID IDW00200101558A patent/ID29273A/id unknown
- 1999-12-01 HK HK02104515.6A patent/HK1043119B/zh not_active IP Right Cessation
- 1999-12-01 US US09/868,704 patent/US6646136B1/en not_active Expired - Lifetime
- 1999-12-01 BR BRPI9916188-5A patent/BR9916188B1/pt not_active IP Right Cessation
- 1999-12-01 JP JP2000588161A patent/JP4790910B2/ja not_active Expired - Lifetime
- 1999-12-01 SK SK816-2001A patent/SK285837B6/sk not_active IP Right Cessation
- 1999-12-01 DK DK99965422T patent/DK1140890T3/da active
- 1999-12-01 DE DE59904407T patent/DE59904407D1/de not_active Expired - Lifetime
- 1999-12-01 WO PCT/EP1999/009333 patent/WO2000035901A1/de not_active Ceased
- 1999-12-01 PT PT99965422T patent/PT1140890E/pt unknown
- 1999-12-01 HU HU0104819A patent/HU229060B1/hu not_active IP Right Cessation
- 1999-12-01 RU RU2001119277/04A patent/RU2233277C2/ru not_active IP Right Cessation
- 1999-12-01 CZ CZ20012145A patent/CZ300544B6/cs not_active IP Right Cessation
- 1999-12-01 EP EP99965422A patent/EP1140890B1/de not_active Expired - Lifetime
- 1999-12-15 TW TW088121987A patent/TWI222866B/zh not_active IP Right Cessation
- 1999-12-17 AR ARP990106530A patent/AR021900A1/es active IP Right Grant
-
2001
- 2001-06-15 NO NO20012964A patent/NO328132B1/no not_active IP Right Cessation
- 2001-07-16 ZA ZA200105840A patent/ZA200105840B/en unknown
Patent Citations (2)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| EP0540914A1 (de) * | 1991-10-28 | 1993-05-12 | Bayer Ag | 2-Aminomethyl-chromane als Wirkstoffe in Arzneimitteln zur Behandlung von Erkrankungen des Zentralnervensystems |
| EP0707007A1 (de) * | 1994-10-14 | 1996-04-17 | MERCK PATENT GmbH | ZNS wirksame Amino(thio)ether-Derivate |
Cited By (1)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| US6903227B2 (en) | 2000-06-02 | 2005-06-07 | Millennium Pharmaceuticals, Inc. | Synthesis of 2-acyl substituted chromanes and intermediates thereof |
Also Published As
Similar Documents
| Publication | Publication Date | Title |
|---|---|---|
| EP1140890B1 (de) | Chromanderivate | |
| DE69307706T2 (de) | Verfahren zur enantioselektiven hydrierung der carbonylgruppe unter verwendung von ruthenium komplexen mit biphosphineliganden | |
| DE10044091A1 (de) | Chromanonderivate | |
| EP1233961B9 (de) | Verfahren zur herstellung von 5-(1-piperazinyl)-benzofuran-2-carboxamid durch übergansmetall-katalysierte aminierung | |
| CH623584A5 (de) | ||
| DE60002303T2 (de) | Verfahren zur herstellung von paroxetin | |
| EP0744401B1 (de) | Verfahren zur Herstellung von optisch aktiven 2-Piperazincarbonsäurederivaten | |
| WO2000026193A1 (de) | Verfahren zur herstellung von 4-[(2',5'-diamino-6'-halogenpyrimidin-4'-yl)amino]-cyclopent-2-enylmethanolen | |
| DE69722565T2 (de) | Verfahren zur Herstellung von 4-Hydroxy-2-Pyrrolidinon und Verfahren zu deren Reinigung | |
| DE69615200T2 (de) | Asymetrische synthese von r und s warfarin und deren ähnlichen verbindungen | |
| EP2812317B1 (de) | Verfahren zur stereoselektiven synthese von 1,4-geschützten 9-hydroxy-5-oxo-1,4-diaza-spiro [5.5]undecane | |
| DE102004033313A1 (de) | Verfahren zur Herstellung von Enantiomeren 3-hydroxy-3-phenyl-propylaminen | |
| EP0551653A1 (de) | Enantiomerenreine 2-Oxa-5,8-diazabicyclo (4.3.0)nonane sowie Verfahren zu ihrer Herstellung | |
| EP0803502B1 (de) | Verfahren zur Herstellung von optisch aktiven 2-Piperazincarbonsäurederivaten | |
| EP1939182A1 (de) | Verfahren zur Herstellung von (R oder S)-2-Alkyl-3-heterocyclyl-1-propanolen | |
| DE4032055A1 (de) | Verfahren zur herstellung von optisch aktiven 4-aryl-2-pyrrolidinonen | |
| EP1125930A1 (de) | Optisch aktive 2-Piperazincarbonsäurederivate und Verfahren zu ihrer Herstellung |
Legal Events
| Date | Code | Title | Description |
|---|---|---|---|
| WWE | Wipo information: entry into national phase |
Ref document number: 99814224.7 Country of ref document: CN |
|
| AK | Designated states |
Kind code of ref document: A1 Designated state(s): AE AL AM AT AU AZ BA BB BG BR BY CA CH CN CU CZ DE DK EE ES FI GB GD GE GH GM HR HU ID IL IN IS JP KE KG KP KR KZ LC LK LR LS LT LU LV MD MG MK MN MW MX NO NZ PL PT RO RU SD SE SG SI SK SL TJ TM TR TT UA UG US UZ VN YU ZA ZW |
|
| AL | Designated countries for regional patents |
Kind code of ref document: A1 Designated state(s): GH GM KE LS MW SD SL SZ TZ UG ZW AM AZ BY KG KZ MD RU TJ TM AT BE CH CY DE DK ES FI FR GB GR IE IT LU MC NL PT SE BF BJ CF CG CI CM GA GN GW ML MR NE SN TD TG |
|
| 121 | Ep: the epo has been informed by wipo that ep was designated in this application | ||
| DFPE | Request for preliminary examination filed prior to expiration of 19th month from priority date (pct application filed before 20040101) | ||
| WWE | Wipo information: entry into national phase |
Ref document number: 1999965422 Country of ref document: EP |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 8162001 Country of ref document: SK |
|
| WWE | Wipo information: entry into national phase |
Ref document number: PV2001-2145 Country of ref document: CZ |
|
| ENP | Entry into the national phase |
Ref document number: 2355098 Country of ref document: CA Ref document number: 2355098 Country of ref document: CA Kind code of ref document: A Ref document number: 2000 588161 Country of ref document: JP Kind code of ref document: A |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 1020017007512 Country of ref document: KR Ref document number: PA/a/2001/006139 Country of ref document: MX |
|
| WWE | Wipo information: entry into national phase |
Ref document number: IN/PCT/2001/733/KOL Country of ref document: IN |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 2001/05840 Country of ref document: ZA Ref document number: 200105840 Country of ref document: ZA Ref document number: 1200100676 Country of ref document: VN |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 20946/00 Country of ref document: AU |
|
| WWP | Wipo information: published in national office |
Ref document number: 1020017007512 Country of ref document: KR |
|
| WWP | Wipo information: published in national office |
Ref document number: PV2001-2145 Country of ref document: CZ |
|
| WWP | Wipo information: published in national office |
Ref document number: 1999965422 Country of ref document: EP |
|
| REG | Reference to national code |
Ref country code: DE Ref legal event code: 8642 |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 09868704 Country of ref document: US |
|
| WWG | Wipo information: grant in national office |
Ref document number: 1999965422 Country of ref document: EP |
|
| WWG | Wipo information: grant in national office |
Ref document number: 20946/00 Country of ref document: AU |
|
| WWG | Wipo information: grant in national office |
Ref document number: 1020017007512 Country of ref document: KR |