WO2001027100A1 - Process for the preparation of 3,4-methylenedioxy-mandelic acid - Google Patents
Process for the preparation of 3,4-methylenedioxy-mandelic acid Download PDFInfo
- Publication number
- WO2001027100A1 WO2001027100A1 PCT/JP2000/007103 JP0007103W WO0127100A1 WO 2001027100 A1 WO2001027100 A1 WO 2001027100A1 JP 0007103 W JP0007103 W JP 0007103W WO 0127100 A1 WO0127100 A1 WO 0127100A1
- Authority
- WO
- WIPO (PCT)
- Prior art keywords
- acid
- methylenedioxybenzene
- producing
- methylenedioxymandelic
- amount
- Prior art date
- Legal status (The legal status is an assumption and is not a legal conclusion. Google has not performed a legal analysis and makes no representation as to the accuracy of the status listed.)
- Ceased
Links
Classifications
-
- C—CHEMISTRY; METALLURGY
- C07—ORGANIC CHEMISTRY
- C07D—HETEROCYCLIC COMPOUNDS
- C07D317/00—Heterocyclic compounds containing five-membered rings having two oxygen atoms as the only ring hetero atoms
- C07D317/08—Heterocyclic compounds containing five-membered rings having two oxygen atoms as the only ring hetero atoms having the hetero atoms in positions 1 and 3
- C07D317/44—Heterocyclic compounds containing five-membered rings having two oxygen atoms as the only ring hetero atoms having the hetero atoms in positions 1 and 3 ortho- or peri-condensed with carbocyclic rings or ring systems
- C07D317/46—Heterocyclic compounds containing five-membered rings having two oxygen atoms as the only ring hetero atoms having the hetero atoms in positions 1 and 3 ortho- or peri-condensed with carbocyclic rings or ring systems condensed with one six-membered ring
- C07D317/48—Methylenedioxybenzenes or hydrogenated methylenedioxybenzenes, unsubstituted on the hetero ring
- C07D317/50—Methylenedioxybenzenes or hydrogenated methylenedioxybenzenes, unsubstituted on the hetero ring with only hydrogen atoms, hydrocarbon or substituted hydrocarbon radicals, directly attached to atoms of the carbocyclic ring
- C07D317/60—Radicals substituted by carbon atoms having three bonds to hetero atoms with at the most one bond to halogen, e.g. ester or nitrile radicals
Definitions
- the present invention relates to a method for producing 3,4-methylenedioxymandelic acid from 1,2-methylenedioxybenzene.
- 3,4-Methylenedioxymandelic acid is a compound useful as a raw material for heliotopin, which is used as a raw material for synthesis of pharmaceuticals and agricultural chemicals and as a general cosmetic flavor.
- a method for producing 3,4-methylenedioxymandelic acid by reacting 1,2-methylenedioxybenzene with glyoxylic acid in the presence of a strong acid such as sulfuric acid or phosphoric acid is already known (for example, Japanese Patent Application Laid-Open No. 54-95573, Perfumer & Flavorist, 14.13 (1989)).
- a strong acid such as sulfuric acid or phosphoric acid
- the produced 3,4-methylenedioxy mandelic acid is insoluble in the present reaction system and thus precipitates as crystals, and as the reaction proceeds, the reaction solution becomes highly viscous or solidifies. Stirring had become extremely difficult.
- JP-A-54-95573 describes that the reaction is preferably carried out without a solvent, but if necessary, addition of formic acid or acetic acid has an advantageous effect on the reaction. Have been. As described above, there is no description of the type of the solvent and its effect, and no specific description is given of the amount of the organic acid when the organic acid is used. Therefore, we conducted intensive studies on this reaction and found that the above problems could be solved by adding an organic solvent. Also, when an organic acid is used, the production rate and selection of 3,4-methylenedioxymandelic acid are unexpectedly improved. It was found that the selectivity was greatly affected by the amount of organic acid added to 1,2-methylenedioxybenzene, and that an optimal amount of organic acid was present.
- An object of the present invention is to react 1,2-methylenedioxybenzene with glyoxylic acid to produce 3,4-methylenedioxymandelic acid with high selectivity. It is intended to provide an industrially suitable method for producing 3,4-methylenedioxymandelic acid, which can be applied even when the content of the product is increased.
- the present invention relates to a method for producing a 3,4-methylenedioxymandelic acid by reacting 1,2-methylenedioxybenzene with glyoxylic acid in the presence of a strong acid, a nonprotonic organic solvent and The reaction is carried out in the presence of at least one selected from the group consisting of 100 to 1200 m1 of organic acid per kg of 1,2-methylenedioxybenzene 3 , 4-Methylenedioxymandelic acid.
- FIG. 1 is a diagram showing an example of a reaction apparatus used for the production method of the present invention.
- the strong acid used in the reaction of the present invention sulfuric acid, inorganic acids such as phosphoric acid; but p K a force 5 '3 following organic acids such as methane sulfonic acid, preferably an inorganic acid, more preferably Sulfuric acid is used. It is preferable to use an aqueous solution of 70% by weight or more of these strong acids.
- the amount used is preferably from 0.5 to 3.00 mol, more preferably from 1.0 to 2.50 mol, per mol of 1,2-methylenedioxybenzene.
- Glyoxylic acid used in the reaction of the present invention a solid (monohydrate) even 4 0 wt 0/0 or more can also be used in aqueous solution.
- the amount used is preferably from 0.8 to 3.0 mol, more preferably from 1.0 to 2.0 mol, per mol of 1,2-methylenedioxybenzene.
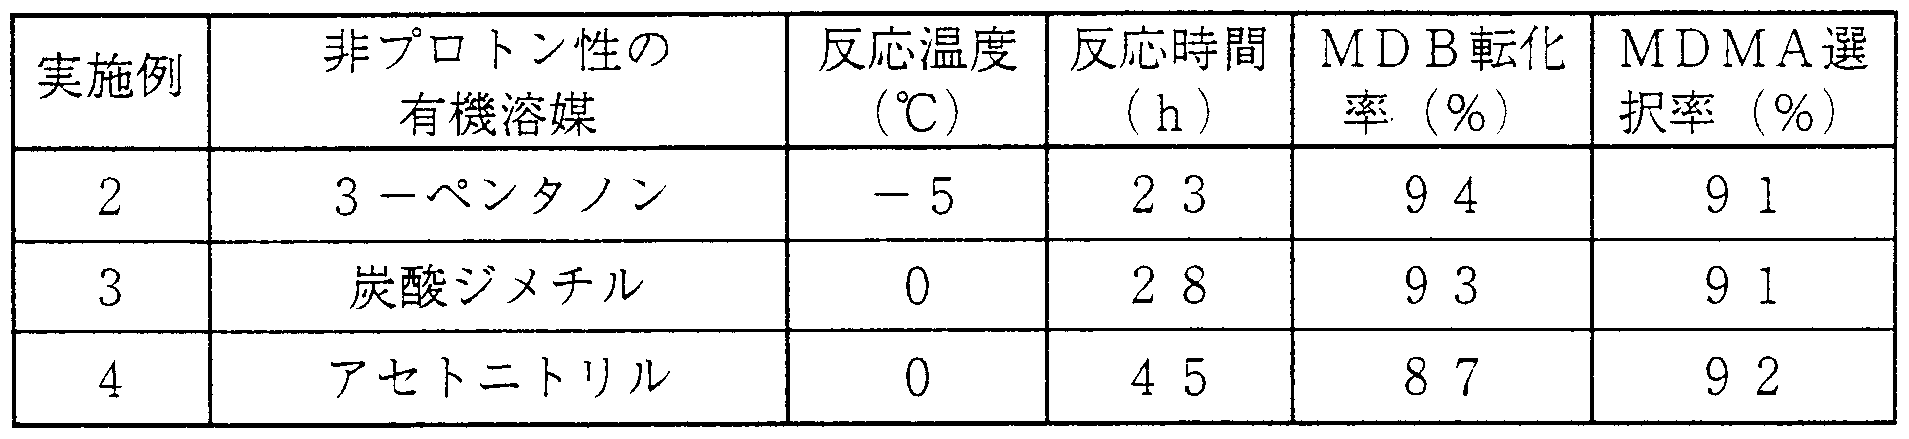
- the aprotic organic solvent used in the reaction of the present invention is not particularly limited as long as it is stable under acidic conditions and does not inhibit the reaction. Examples thereof include getyl ether, diisopropyl ether, dibutyl ether, and tetrahydrofuran.
- Ethers such as acetone; ketones such as acetone, 2-butanone, 2-pentanone, 3-pentanone, 4-methyl-12-pentanone, cyclopentanone, cyclohexanone; ethyl formate, isopropyl formate, butyl formate
- Carboxylate esters such as acetic acid, methyl acetate, ethyl acetate, isopropyl acetate, butyl acetate, methyl propionate, ethyl propionate, isopropyl propionate and butyl propionate; N, N-dimethylformamide, N, N —Dimethylacetamide, Hexamethylphosphorite Amides such as triamide and 1-methyl-12-pyrrolidone; ureas such as 1,3-dimethyl-2-imidazolidone; carbonates such as dimethyl carbonate and getyl carbonate; sulfoxides such as dimethyl sulfoxide.
- the amount of the aprotic organic solvent to be used is preferably 100 to 2000 ml, more preferably 100 to 1000 ml, per 1 kg of 1,2-methylenedioxybenzene. These organic solvents may be used alone or in combination of two or more.
- the organic acid used in the reaction of the present invention is not particularly limited as long as it is stable under acidic conditions and does not inhibit the reaction.
- aliphatic carboxylic acids such as trifluoroacetic acid and dichloroacetic acid, preferably aliphatic carboxylic acids, and more preferably acetic acid.
- These organic acids may be used alone or in combination of two or more.
- the amount of the organic acid to be used is 100 to 1200 ml, preferably 100 to 1000 ml, more preferably 100 to 500 m per 1 kg of 1,2-methylenedioxybenzene. Is one. If the amount used is less than 100 ml, the agitation of the reaction solution will be extremely poor, and the temperature of the solution will be partially increased, by-products will be generated, and the selectivity of the target product will be reduced. On the other hand, when it is more than 120 Om 1, the reaction rate becomes extremely slow, and there is a problem that the reaction takes a long time and the selectivity of the target product is lowered.
- the reaction is usually carried out under normal pressure. If necessary, the reaction may be carried out under increased or reduced pressure.
- the reaction time is generally 1 to 100 hours, preferably about 3 to 75 hours.
- the product obtained is neutralized by adding an appropriate amount of base after the reaction is completed, extracted with an appropriate solvent, and separated and purified by a general method such as column chromatography, distillation, or recrystallization. Is done.
- the reaction was carried out in the same manner as in Example 2 except that no organic solvent was added. Then, the reaction solution became highly viscous during the reaction, and only the periphery of the stirring spring was stirred, and the whole could not be sufficiently stirred. As a result, the conversion of 1,2-methylenedioxybenzene was 94%, and the selectivity of 3,4-methylenedioxymandelic acid was 77%.
- Example 6 was repeated in the same manner as in Example 6, except that the amount of acetic acid used for 1,2-methylenedioxybenzene was outside the scope of the present invention and the reaction temperature and reaction time were changed. The reaction was performed. The results are shown in Table 2 .: Table 2
- 3,4-methylenedioxymandelic acid can be produced with high selectivity by reacting 1,2-methylenedioxybenzene with glyoxylic acid. It is possible to provide an industrially suitable method for producing 3,4-methylenedioximandelic acid, which can be applied even when it is raised.
Landscapes
- Chemical & Material Sciences (AREA)
- Organic Chemistry (AREA)
- Organic Low-Molecular-Weight Compounds And Preparation Thereof (AREA)
- Low-Molecular Organic Synthesis Reactions Using Catalysts (AREA)
Description

Claims
Priority Applications (3)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| US10/110,263 US6603024B1 (en) | 1999-10-13 | 2000-10-13 | Process for the preparation of 3,4-methylenedioxymandelic acid |
| EP00966457A EP1229031B1 (en) | 1999-10-13 | 2000-10-13 | Process for the preparation of 3,4-methylenedioxy-mandelic acid |
| NO20021727A NO327178B1 (no) | 1999-10-13 | 2002-04-12 | Fremgangsmate for fremstilling av 3,4-metylendioksy-mandelsyre |
Applications Claiming Priority (4)
| Application Number | Priority Date | Filing Date | Title |
|---|---|---|---|
| JP29077499 | 1999-10-13 | ||
| JP11/290774 | 1999-10-13 | ||
| JP11/326116 | 1999-11-16 | ||
| JP32611699 | 1999-11-16 |
Publications (1)
| Publication Number | Publication Date |
|---|---|
| WO2001027100A1 true WO2001027100A1 (en) | 2001-04-19 |
Family
ID=26558227
Family Applications (1)
| Application Number | Title | Priority Date | Filing Date |
|---|---|---|---|
| PCT/JP2000/007103 Ceased WO2001027100A1 (en) | 1999-10-13 | 2000-10-13 | Process for the preparation of 3,4-methylenedioxy-mandelic acid |
Country Status (7)
| Country | Link |
|---|---|
| US (1) | US6603024B1 (ja) |
| EP (1) | EP1229031B1 (ja) |
| KR (1) | KR100721598B1 (ja) |
| CN (1) | CN1220689C (ja) |
| NO (1) | NO327178B1 (ja) |
| TW (1) | TWI226888B (ja) |
| WO (1) | WO2001027100A1 (ja) |
Families Citing this family (7)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| TW200819420A (en) * | 2006-10-31 | 2008-05-01 | Univ Nat Chunghsing | The alkoxylation method applied to extend phenol compounds to have alkoxy functional group |
| TW200819421A (en) * | 2006-10-31 | 2008-05-01 | Univ Nat Chunghsing | The method of synthesizing biphenol A, BPA having di-alkoxyl group by using polycarbonate or its waste |
| CN104151285A (zh) * | 2014-08-14 | 2014-11-19 | 江苏亚泰化工有限公司 | 一种制备3,4-亚甲二氧基杏仁酸的方法 |
| GR1008653B (el) * | 2014-12-23 | 2016-01-18 | ΦΑΡΜΑΤΕΝ ΑΝΩΝΥΜΟΣ ΒΙΟΜΗΧΑΝΙΚΗ ΚΑΙ ΕΜΠΟΡΙΚΗ ΕΤΑΙΡΕΙΑ ΦΑΡΜΑΚΕΥΤΙΚΩΝ ΙΑΤΡΙΚΩΝ ΚΑΙ ΚΑΛΛΥΝΤΙΚΩΝ ΠΡΟΪΟΝΤΩΝ με δ.τ. ΦΑΡΜΑΤΕΝ, | Μεθοδος δια την παρασκευη πιπεροναλης |
| CN105837552A (zh) * | 2016-04-01 | 2016-08-10 | 衢州信步化工科技有限公司 | 一种洋茉莉醛中间体合成反应液的处理方法 |
| CN105777705B (zh) * | 2016-04-01 | 2018-09-21 | 福建仁宏医药化工有限公司 | 一种洋茉莉醛中间体合成反应后期处理方法 |
| CN110684008B (zh) * | 2018-07-04 | 2023-01-20 | 天津大学 | 一种乳化工艺合成3,4-亚甲二氧基苯乙醇酸的方法及装置 |
Citations (2)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| EP0002460A1 (de) * | 1977-12-07 | 1979-06-27 | Haarmann & Reimer Gmbh | Verfahren zur Herstellung von 3,4-Methylendioxymandelsäure |
| JPH0859650A (ja) * | 1994-08-12 | 1996-03-05 | Ube Ind Ltd | 3,4−メチレンジオキシマンデル酸の製造方法 |
Family Cites Families (2)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| DE2824407A1 (de) | 1978-06-03 | 1979-12-13 | Bayer Ag | Verfahren zur herstellung von arylglyoxylsaeuren |
| CA2238976A1 (en) * | 1995-12-12 | 1997-06-19 | Merck & Co., Inc. | Process for the preparation of an endothelin antagonist |
-
2000
- 2000-10-04 TW TW089120625A patent/TWI226888B/zh not_active IP Right Cessation
- 2000-10-13 US US10/110,263 patent/US6603024B1/en not_active Expired - Fee Related
- 2000-10-13 WO PCT/JP2000/007103 patent/WO2001027100A1/ja not_active Ceased
- 2000-10-13 CN CNB008142203A patent/CN1220689C/zh not_active Expired - Fee Related
- 2000-10-13 EP EP00966457A patent/EP1229031B1/en not_active Expired - Lifetime
- 2000-10-13 KR KR1020027004652A patent/KR100721598B1/ko not_active Expired - Fee Related
-
2002
- 2002-04-12 NO NO20021727A patent/NO327178B1/no not_active IP Right Cessation
Patent Citations (2)
| Publication number | Priority date | Publication date | Assignee | Title |
|---|---|---|---|---|
| EP0002460A1 (de) * | 1977-12-07 | 1979-06-27 | Haarmann & Reimer Gmbh | Verfahren zur Herstellung von 3,4-Methylendioxymandelsäure |
| JPH0859650A (ja) * | 1994-08-12 | 1996-03-05 | Ube Ind Ltd | 3,4−メチレンジオキシマンデル酸の製造方法 |
Non-Patent Citations (1)
| Title |
|---|
| See also references of EP1229031A4 * |
Also Published As
| Publication number | Publication date |
|---|---|
| CN1220689C (zh) | 2005-09-28 |
| EP1229031A1 (en) | 2002-08-07 |
| TWI226888B (en) | 2005-01-21 |
| EP1229031B1 (en) | 2004-04-28 |
| KR20020068520A (ko) | 2002-08-27 |
| CN1379770A (zh) | 2002-11-13 |
| KR100721598B1 (ko) | 2007-05-25 |
| NO20021727L (no) | 2002-04-12 |
| US6603024B1 (en) | 2003-08-05 |
| EP1229031A4 (en) | 2003-04-16 |
| NO327178B1 (no) | 2009-05-04 |
| NO20021727D0 (no) | 2002-04-12 |
Similar Documents
| Publication | Publication Date | Title |
|---|---|---|
| JP4186404B2 (ja) | 3,4−メチレンジオキシマンデル酸の製造方法 | |
| JP2006160663A (ja) | 1,1’−ビス(2−ヒドロキシナフチル)類の製造方法 | |
| US6603024B1 (en) | Process for the preparation of 3,4-methylenedioxymandelic acid | |
| JP2001206883A (ja) | 3,4−メチレンジオキシマンデル酸の製造法 | |
| JP4186405B2 (ja) | 3,4−メチレンジオキシマンデル酸の製法 | |
| KR101940994B1 (ko) | 4‘-포르밀-4-비페닐카르복실산의 신규 제조방법 | |
| CN108084093B (zh) | 一锅法合成3-二氟甲基-1-甲基-1h-吡唑-4-羧酸的方法 | |
| JPH0717935A (ja) | 3−アミノプロピオニトリルの製造法 | |
| JPH021819B2 (ja) | ||
| JP2005336101A (ja) | フェニルオキサジアゾール類の製造法 | |
| JPH053859B2 (ja) | ||
| KR101393010B1 (ko) | 템포 촉매제를 이용한 2-(4-포르밀페닐)프로피온 산의 제조방법 | |
| EP0468727B1 (en) | Process for producing hydrazone derivative | |
| JP4427661B2 (ja) | 2−アルキリデン−4−ブロモアセト酢酸エステル類の製造方法 | |
| JPS60501709A (ja) | シアノヒドリンの製造方法 | |
| JPH03258768A (ja) | 1,3,5―パーヒドロトリアジン―2,4,6―トリチオン誘導体の製造方法 | |
| JP2001097937A (ja) | 4−フルオロ−2−トリフルオロメチルベンゾニトリルの製造方法 | |
| WO2002096853A1 (fr) | Procede de production d'un derive d'aralkylamine | |
| JPS5883683A (ja) | フエナジン−5−オキシドの製造法 | |
| JPH10182544A (ja) | p−アシルアルキルベンゼン誘導体の製造法 | |
| JP2002173459A (ja) | 2,4,5−トリアルキルベンズアルデヒドの製造方法 | |
| JPH08119960A (ja) | ビスラクトンの製造方法 | |
| JPH11279169A (ja) | 3−イソクロマノン類の製造方法 | |
| JPH04316550A (ja) | シアノ酢酸エステルの製造法 | |
| JPH1067764A (ja) | 3−メチルテトラヒドロフランを製造する方法 |
Legal Events
| Date | Code | Title | Description |
|---|---|---|---|
| AK | Designated states |
Kind code of ref document: A1 Designated state(s): CN IN KR NO US |
|
| AL | Designated countries for regional patents |
Kind code of ref document: A1 Designated state(s): AT BE CH CY DE DK ES FI FR GB GR IE IT LU MC NL PT SE |
|
| DFPE | Request for preliminary examination filed prior to expiration of 19th month from priority date (pct application filed before 20040101) | ||
| 121 | Ep: the epo has been informed by wipo that ep was designated in this application | ||
| WWE | Wipo information: entry into national phase |
Ref document number: IN/PCT/2002/00329/DE Country of ref document: IN |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 10110263 Country of ref document: US |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 1020027004652 Country of ref document: KR |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 008142203 Country of ref document: CN |
|
| WWE | Wipo information: entry into national phase |
Ref document number: 2000966457 Country of ref document: EP |
|
| WWP | Wipo information: published in national office |
Ref document number: 2000966457 Country of ref document: EP |
|
| WWP | Wipo information: published in national office |
Ref document number: 1020027004652 Country of ref document: KR |
|
| WWG | Wipo information: grant in national office |
Ref document number: 2000966457 Country of ref document: EP |